Journal Description
Thalassemia Reports
Thalassemia Reports
is an international, peer-reviewed, open access journal on the study, diagnosis, and treatment of thalassemia, published quarterly online by MDPI (from Volume 12, Issue 1 - 2022).
- Open Access— free for readers, with article processing charges (APC) paid by authors or their institutions.
- High Visibility: indexed within ESCI (Web of Science), Embase, and other databases.
- Journal Rank: JCR - Q2 (Hematology)
- Rapid Publication: manuscripts are peer-reviewed and a first decision is provided to authors approximately 40.5 days after submission; acceptance to publication is undertaken in 8.7 days (median values for papers published in this journal in the first half of 2026).
- Recognition of Reviewers: Reviewers whose reports are timely and of high quality receive an APC discount voucher for a future publication in an MDPI journal. Become a reviewer.
- Journal Clusters of Hematology: Hemato, Hematology Reports, Thalassemia Reports and Journal of Clinical Medicine.
Impact Factor:
2.4 (2025);
5-Year Impact Factor:
1.6 (2025)
Latest Articles
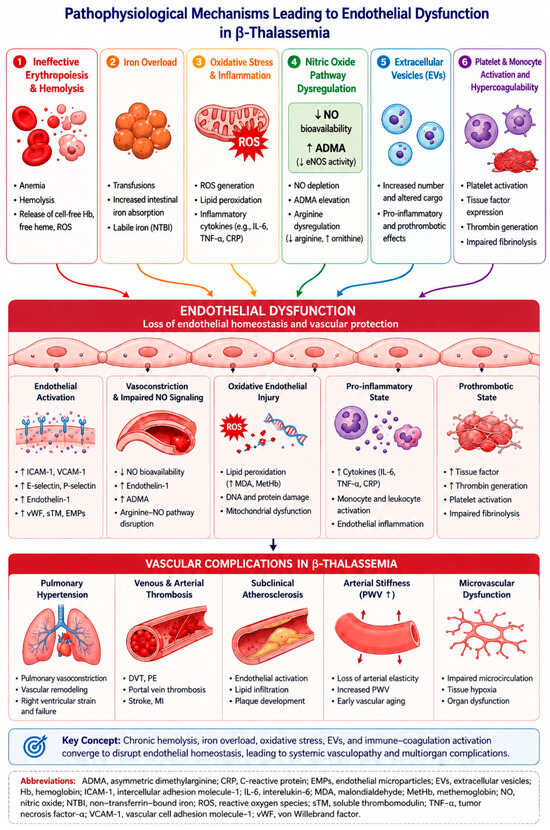
Endothelial Dysfunction in β-Thalassemia: Mechanisms, Biomarkers, Vascular Imaging and Therapeutic Perspectives
Thalass. Rep. 2026, 16(3), 22; https://doi.org/10.3390/thalassrep16030022 - 7 Sep 2026
Abstract
Background/Objectives: Endothelial dysfunction is increasingly recognized as a key contributor to vascular complications in β-thalassemia but remains insufficiently integrated into clinical assessment and therapeutic trial design. This review summarizes current evidence on the mechanisms, biomarkers, vascular imaging findings, and therapeutic perspectives of endothelial
[...] Read more.
Background/Objectives: Endothelial dysfunction is increasingly recognized as a key contributor to vascular complications in β-thalassemia but remains insufficiently integrated into clinical assessment and therapeutic trial design. This review summarizes current evidence on the mechanisms, biomarkers, vascular imaging findings, and therapeutic perspectives of endothelial dysfunction across the clinical spectrum of β-thalassemia. Methods: A focused PubMed search was conducted using predefined terms related to endothelial dysfunction, nitric oxide, oxidative stress, adhesion molecules, extracellular vesicles, vascular imaging, and therapeutics in β-thalassemia. Original studies and systematic reviews evaluating endothelial dysfunction, vascular biomarkers, imaging, or cardiovascular outcomes in β-thalassemia were included. Results: Endothelial dysfunction arises from the interplay of hemolysis-driven nitric oxide depletion, free heme- and iron-mediated oxidative injury, chronic inflammation, extracellular vesicle-mediated vascular activation, and hypercoagulability. Meta-analyses demonstrate significant elevations in intercellular adhesion molecule-1, vascular cell adhesion molecule-1, E-selectin, P-selectin, and endothelin-1. Emerging biomarkers include endothelial activation and stress index, adipocytokines, and serum metabolomics. Vascular imaging demonstrates impaired flow-mediated dilation, increased carotid intima-media thickness, elevated pulse-wave velocity, and pulmonary hypertension, with distinct vascular phenotypes in transfusion-dependent thalassemia (TDT) and non-transfusion-dependent thalassemia (NTDT). Iron chelation, luspatercept, mitapivat, and curative gene-based therapies have strong biological rationale for improving endothelial function, although vascular endpoints have rarely been evaluated. Conclusions: Endothelial dysfunction is a multidimensional and potentially modifiable component of β-thalassemia vasculopathy. Future clinical trials should incorporate standardized endothelial biomarkers and vascular imaging as predefined endpoints and evaluate TDT and NTDT separately to advance mechanism-based vascular therapies.
Full article
(This article belongs to the Section Innovative Treatment of Thalassemia)
►
Show Figures
Open AccessArticle
Circulating STAT3 and SMAD1/SMAD4 with Hepcidin Levels Across β-Thalassemia Subtypes: A Cross-Sectional Study
by
Kirti Upadhyay, Nitu Nigam, Swasti Sinha, Nishant Verma, Suresh Chand, Anshul Gupta and Soniya Nityanand
Thalass. Rep. 2026, 16(3), 21; https://doi.org/10.3390/thalassrep16030021 - 3 Sep 2026
Abstract
Background/Objectives: The background of the research shows that β-thalassemia is a hemoglobinopathy which is characterized by ineffective erythropoiesis and iron overload and this is kept in balance by hepcidin hormone due to BMP/SMAD and JAK/STAT3 pathways. However, it is not known how exactly
[...] Read more.
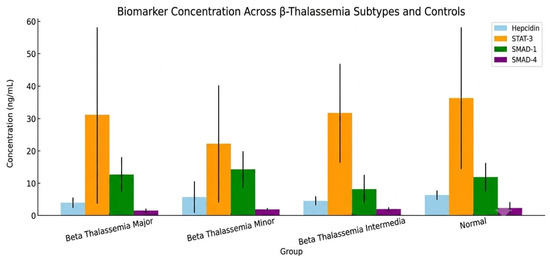
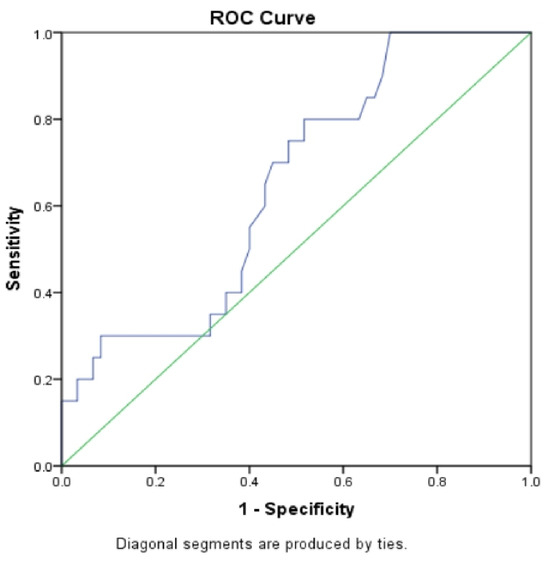
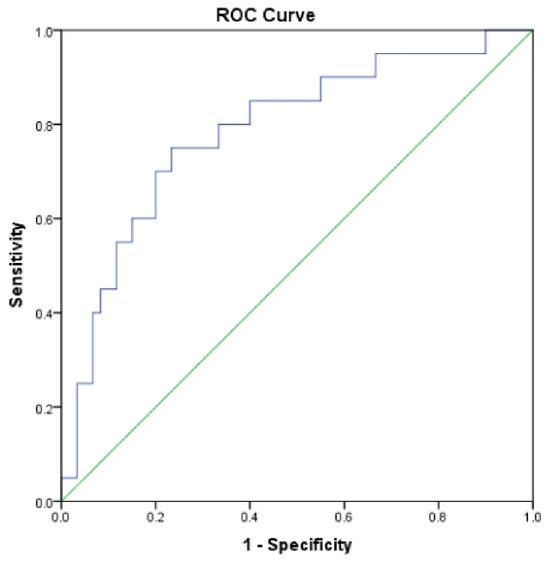
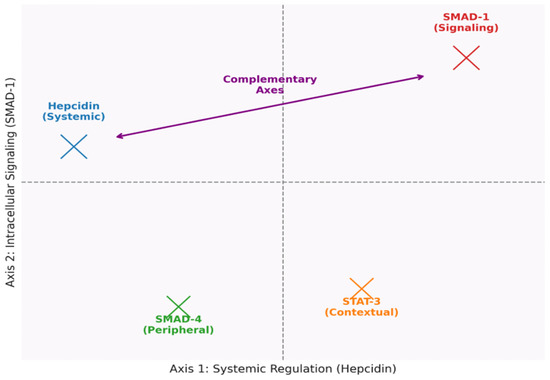
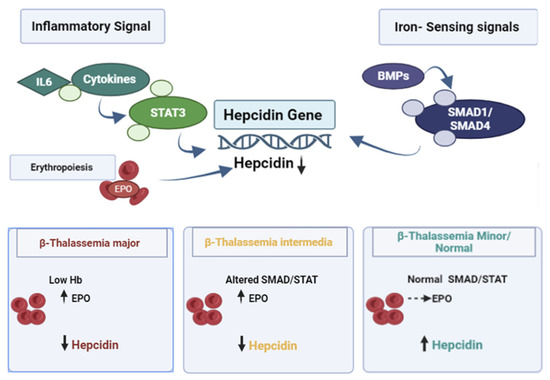
Background/Objectives: The background of the research shows that β-thalassemia is a hemoglobinopathy which is characterized by ineffective erythropoiesis and iron overload and this is kept in balance by hepcidin hormone due to BMP/SMAD and JAK/STAT3 pathways. However, it is not known how exactly these signaling pathways are involved in iron metabolism disorders for different forms of β-thalassemia, therefore, the aim of the present research was to analyze circulating levels of hepcidin, STAT3, SMAD1 and SMAD4 in participants with β-thalassemia and evaluate their exploratory discriminatory potential. Methods: The methodology of the research includes a comparative cross-sectional study of 135 patients with β-thalassemia major, intermedia and minor, as well as healthy control individuals based on hemoglobinopathy typing with the high-performance liquid chromatography. The circulating hepcidin levels, SMAD1 and SMAD4 concentrations as well as plasma STAT3 levels were identified and measured by ELISA method. Statistical processing methods include one-way ANOVA, multinomial logistic regression and ROC curve analysis based on ten-fold cross-validation. Results: Hepcidin and SMAD1 levels showed heterogeneity among groups, whereas differences in STAT3 and SMAD4 were not statistically significant. In multinomial logistic regression, lower hepcidin was associated with β-thalassemia major and lower SMAD1 with β-thalassemia intermedia. ROC analysis showed moderate in-sample discrimination for hepcidin and stronger discrimination for SMAD1. Conclusions: Altered circulating hepcidin and SMAD1 patterns may reflect dysregulation of BMP-SMAD/hepcidin signaling across β-thalassemia subtypes. These findings are exploratory and require independent validation before clinical application.
Full article
(This article belongs to the Section Innovative Treatment of Thalassemia)
►▼
Show Figures

Figure 1
Open AccessArticle
Exercise and Physical Activity in Patients with Sickle Cell Disease in Jeddah, Saudi Arabia: A Cross-Sectional Study
by
Ahmed S. Barefah, Hatem M. Alahwal, Eman M. Mansory, Ahmed H. Alabdele, Naif N. Althagafi, Raad M. Alsaadi, Ahmed O. Qumsani, Abdullah H. Bokhary, Nasser F. Alshanbari, Abdulrahman S. Almaghrabi, Khalid S. Basahih, Osman O. Radhwi and Salem M. Bahashwan
Thalass. Rep. 2026, 16(3), 20; https://doi.org/10.3390/thalassrep16030020 - 2 Sep 2026
Abstract
►▼
Show Figures
Background: Sickle cell disease (SCD) is a hereditary hemoglobinopathy characterized by chronic anemia, recurrent pain episodes, and multi-organ complications, which collectively impact patients’ quality of life and functional capacity. Exercise and physical activity are recognized as essential components of general health and well-being;
[...] Read more.
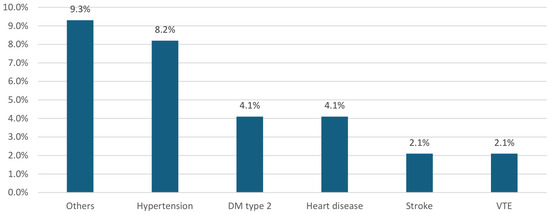
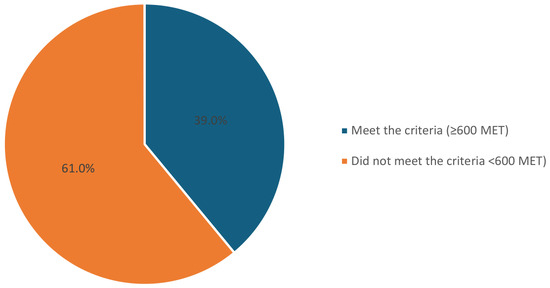
Background: Sickle cell disease (SCD) is a hereditary hemoglobinopathy characterized by chronic anemia, recurrent pain episodes, and multi-organ complications, which collectively impact patients’ quality of life and functional capacity. Exercise and physical activity are recognized as essential components of general health and well-being; however, their role in individuals with SCD remains underexplored due to concerns regarding safety, tolerance, and the potential to precipitate vaso-occlusive crises. This study aimed to assess the patterns and frequency of exercise and physical activity among patients with sickle cell disease using a validated questionnaire. Methods: We conducted a cross-sectional study on patients diagnosed with sickle cell disease at King Abdulaziz University Hospital in Jeddah, Saudi Arabia. After obtaining ethical approval, we contacted the patients by phone or met them in person and administered the Global Physical Activity Questionnaire (GPAQ). Data on age, gender, comorbid conditions, disease-related variables, and weekly physical activity were collected. Total physical activity was expressed in metabolic equivalent of task (MET)-minutes per week, and participants achieving ≥600 MET-minutes per week were classified as meeting the World Health Organization (WHO) recommendation. Results: Of the 97 patients, males represented 52.6% of the participants, with a median age of 33 years. Overall, 70.1% of the patients were on hydroxyurea, and 14.4% had undergone splenectomy. More than half of the patients (56.7%) reported difficulties with exercise. According to the findings, 39% of the patients were getting enough physical activity each week, as recommended by the WHO. Male patients were significantly more likely to meet the WHO physical activity recommendation (p = 0.003), whereas no statistically significant associations were observed for age, nationality, body mass index, hemoglobin level, hydroxyurea use, splenectomy, hospital admissions, transfusion history, comorbidities, or exercise-related symptoms (all p > 0.05). Conclusions: Our data suggest that a significant proportion of patients with SCD did not achieve the WHO-recommended physical activity level, with adherence observed in only about forty percent of participants, and that adherence was more common in male patients (adjusted odds ratio 3.30, 95% CI 1.36–8.52, after multivariable adjustment for age, body mass index, hemoglobin, and transfusion status). This association was concentrated in occupational rather than recreational activity and was not explained by the clinical variables measured. Because no comparison group was studied, the extent to which this shortfall is attributable to SCD itself, rather than to the population-level inactivity already documented in Saudi Arabia, cannot be determined from these data. These findings identify a potential area for targeted intervention, particularly among women, and underscore the need for further research into disease-specific and sociocultural barriers before tailored programs can be developed. Further larger, controlled, multi-center studies are necessary to clarify the specific contribution of SCD to inactivity and to guide individualized physical activity promotion in this population.
Full article

Figure 1
Open AccessArticle
An Integrated Gene Therapy Strategy for γ-Globin Addition and HbF Reactivation in β-Thalassemia
by
Kiriaki Paschoudi, Ninos Ioannis Vasiloudis, Fotios Papadopoulos, Xenia Nikolaou, Anastasia Papadopoulou, Pavel Sova, Evangelia Yannaki and Nikoletta Psatha
Thalass. Rep. 2026, 16(3), 19; https://doi.org/10.3390/thalassrep16030019 - 1 Sep 2026
Abstract
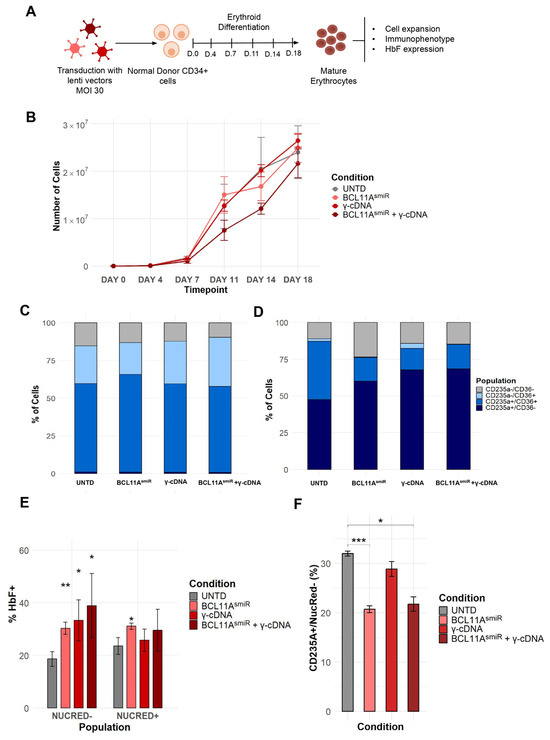
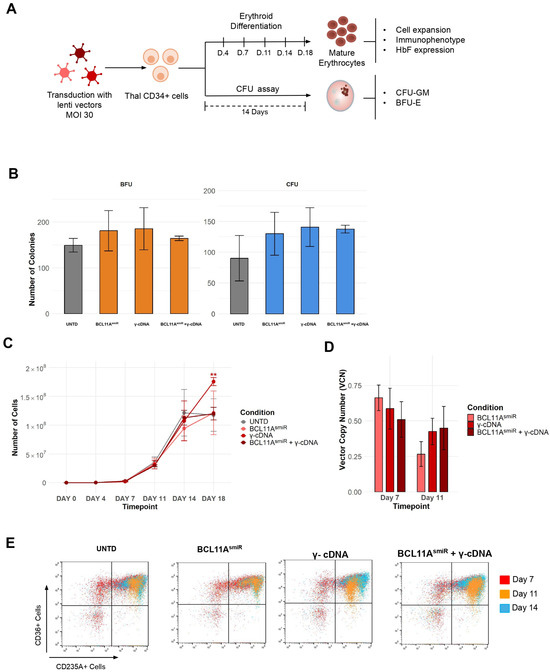
Background: β-Thalassemia and other β-hemoglobinopathies arise from defective β-globin production, and reactivating fetal hemoglobin (HbF) is a well-established strategy to ameliorate disease severity. Two principal gene-therapy approaches—erythroid-specific γ-globin gene addition and shRNA/shmiRNA-mediated knockdown of the γ-globin repressor BCL11A—have each shown clinical promise, but
[...] Read more.
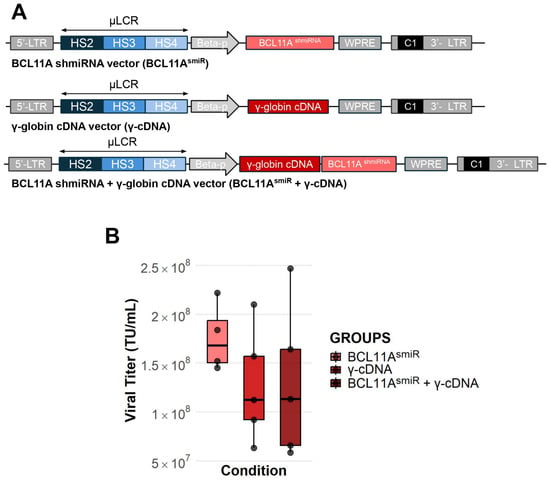
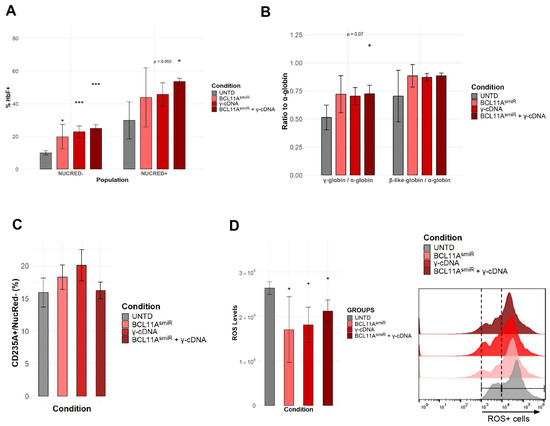
Background: β-Thalassemia and other β-hemoglobinopathies arise from defective β-globin production, and reactivating fetal hemoglobin (HbF) is a well-established strategy to ameliorate disease severity. Two principal gene-therapy approaches—erythroid-specific γ-globin gene addition and shRNA/shmiRNA-mediated knockdown of the γ-globin repressor BCL11A—have each shown clinical promise, but their combination within a single vector remains largely unexplored. Methods: Here we developed and compared three compact, C1-insulated lentiviral vectors driven by the micro-locus control region (μLCR): a BCL11A-targeting shmiRNA vector, a γ-globin cDNA vector, and a dual construct combining both elements. Results: All vectors were produced at high titers, and incorporating both cassettes into a single construct did not compromise vector production or cell viability. In CD34+ hematopoietic stem and progenitor cells from healthy donors and β-thalassemia patients, transduction did not impair proliferation, erythroid differentiation, or progenitor colony formation. All three vectors significantly increased the proportion of HbF-expressing cells, with the combined shmiRNA + γ-globin cDNA vector achieving the highest induction, reaching approximately 25% HbF-positive cells in the enucleated erythroid population of thalassemic cells versus approximately 10% in untransduced controls. Notably, all constructs significantly reduced reactive oxygen species levels, indicating alleviation of oxidative stress. Conclusions: Together, these findings support the feasibility and therapeutic potential of combining complementary HbF-inducing mechanisms within a single lentiviral vector for β-thalassemia.
Full article
(This article belongs to the Section Innovative Treatment of Thalassemia)
►▼
Show Figures

Figure 1
Open AccessReview
Genetic Approach in Diagnosis and Follow-Up of Patients with Thalassemia: A Comprehensive Narrative Review
by
Ashraf T. Soliman, Fawzia Alyafei, Nada Alaaraj, Noor Hamed, Shayma Ahmed and Ahmed Elawwa
Thalass. Rep. 2026, 16(3), 18; https://doi.org/10.3390/thalassrep16030018 - 19 Aug 2026
Abstract
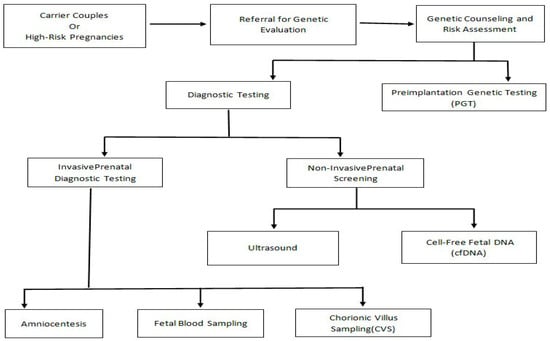
Thalassemia represents the world’s most prevalent inherited hemoglobin disorder, affecting approximately 4.4 per 10,000 live births globally. Accurate genetic characterization is indispensable both for definitive diagnosis and for lifetime clinical monitoring. The past two decades have witnessed a paradigm shift from conventional protein-based
[...] Read more.
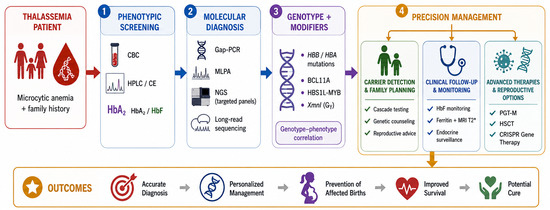
Thalassemia represents the world’s most prevalent inherited hemoglobin disorder, affecting approximately 4.4 per 10,000 live births globally. Accurate genetic characterization is indispensable both for definitive diagnosis and for lifetime clinical monitoring. The past two decades have witnessed a paradigm shift from conventional protein-based assays toward comprehensive molecular techniques, including next-generation sequencing (NGS) and third-generation (long-read) sequencing, which in turn have enabled reproductive applications such as preimplantation genetic testing for monogenic disease (PGT-M) to identify unaffected embryos before implantation. (1) To systematically evaluate the molecular techniques available for confirming the diagnosis of alpha- and beta-thalassemia, including their diagnostic accuracy, indications, and limitations; (2) to examine how genotype–phenotype correlation and genetic modifier profiling inform clinical prognosis and therapeutic decision-making; and (3) to define evidence-based genetic monitoring parameters for longitudinal follow-up of patients receiving transfusions, iron chelation, and novel curative therapies including gene therapy. A comprehensive narrative review was conducted by systematically searching PubMed/MEDLINE for English-language peer-reviewed articles published between January 2000 and December 2024. Forty-three studies were ultimately included after applying predefined inclusion and exclusion criteria. Quality of included studies was assessed using SANRA (Scale for the Assessment of Narrative Review Articles). HPLC and capillary electrophoresis remain first-line phenotyping tools; DNA-based confirmation is mandatory for complete genotyping. Among known, previously characterized mutations, NGS-based targeted panels achieve > 95% detection sensitivity, but they require MLPA co-testing or long-read sequencing to detect structural variants such as large deletions. Genotype–phenotype prediction is substantially improved, though not rendered fully deterministic, by profiling three major modifier loci: XmnI (Gγ), BCL11A, and HBS1L-MYB. PGT-M using NGS achieves near-complete genotyping accuracy (>99%) with live birth rates of 40–60% per frozen embryo transfer cycle. For patients receiving curative gene therapy (exagamglogene autotemcel/Casgevy), molecular follow-up protocols spanning 15 years are now recommended. Cardiac T2* MRI remains the most reliable non-invasive tool for iron overload follow-up, superior to serum ferritin alone. A tiered, genotype-informed approach—combining HPLC/CE phenotyping, targeted molecular diagnostics, genetic modifier profiling, and periodic re-evaluation—optimizes diagnostic precision and guides individualized management across the thalassemia spectrum. Integration of PGT-M and long-read sequencing into standard care pathways, alongside robust gene therapy follow-up protocols, will define the next era of thalassemia genetics.
Full article
(This article belongs to the Special Issue Genetic Approach in Diagnosis and Following Up of Patients With Thalassemia)
►▼
Show Figures

Graphical abstract
Open AccessArticle
The Role of Thalassaemic Red Cells in the Mechanism Process of Hypercoagulable State in Thalassaemia
by
Wardah Roslan, Rosnah Bahar, Mohd Nazri Hassan, Norsarwany Mohamad, Shafini Mohamed Yusoff, Salfarina Iberahim, Zefarina Zulkafli, Noor Haslina Mohd Noor, Marini Ramli, Razan Hayati Zulkeflee, Marne Abdullah and Wan Suriana Wan Ab Rahman
Thalass. Rep. 2026, 16(3), 17; https://doi.org/10.3390/thalassrep16030017 - 18 Aug 2026
Abstract
Background: Thromboembolic events (TEEs) are recognized complications in thalassaemia, arising from a hypercoagulable state driven by multiple proposed mechanisms. These concerns have prompted numerous investigations to better understand the underlying prothrombotic pathways. This study aimed to compare and correlate the levels of fragmented
[...] Read more.
Background: Thromboembolic events (TEEs) are recognized complications in thalassaemia, arising from a hypercoagulable state driven by multiple proposed mechanisms. These concerns have prompted numerous investigations to better understand the underlying prothrombotic pathways. This study aimed to compare and correlate the levels of fragmented red blood cells (FRCs), hypercoagulable protein markers, and erythrocyte phosphatidylserine (PS) exposure in thalassaemia patients. This 12-month prospective case-control study at HPUSM recruited 34 thalassaemia patients (21 with thalassaemia major and 13 with thalassaemia intermedia) and 10 healthy controls. Methods: FRCs, erythrocyte PS, protein C, free protein S, and antithrombin III levels were measured. Results: A total of 44 subjects were included, with a mean age of 22.43 ± 10.45 years; most were male (61.4%) and Malay (86.4%). Median FRCs and PS exposure were significantly elevated in both thalassaemia groups compared to controls (p < 0.001 for FRCs; p = 0.048 for PS). Mean protein C and free protein S levels were significantly reduced in thalassaemia major (55.00 ± 10.20% and 65.77 ± 8.66%) and thalassaemia intermedia (61.23 ± 16.99% and 61.11 ± 14.65%) compared to controls (101.60 ± 18.97% and 95.12 ± 23.57%) (p < 0.001 for both), while antithrombin III levels were not significantly different. A significant negative correlation was observed between PS exposure and protein C (p = 0.014). Protein C was positively correlated with free protein S (p = 0.021) and antithrombin III (p < 0.001). Conclusions: Elevated PS exposure and reduced protein C and free protein S levels in thalassaemia patients indicate a potential procoagulant phenotype.
Full article
Open AccessCase Report
A Novel α-Globin Gene Variant: Hb Romagna [α61(E10)Lys>Gln; HBA1:c.184A>C] Co-Inherited with Hb A2-Lampang [δ47(CD6)Asp>Asn; HBD:c.142G>A] in an Italian Diabetic Woman
by
Marco Rosetti, Giovanni Poletti, Melania Olivieri, Massimo Mogni, Massimo Maffei, Abbate Noemi, Pisani Raffaele, Sauro Maoggi, Domenico Coviello and Giovanni Ivaldi
Thalass. Rep. 2026, 16(3), 16; https://doi.org/10.3390/thalassrep16030016 - 4 Aug 2026
Abstract
Background: Screening for hemoglobinopathies, particularly in Mediterranean countries, is primarily aimed at preventing beta-thalassemia. However, during such screening, numerous other Hb defects can be diagnosed, each with different clinical significance, including qualitative and quantitative variations in the alpha genes. Aims: In the present
[...] Read more.
Background: Screening for hemoglobinopathies, particularly in Mediterranean countries, is primarily aimed at preventing beta-thalassemia. However, during such screening, numerous other Hb defects can be diagnosed, each with different clinical significance, including qualitative and quantitative variations in the alpha genes. Aims: In the present study, we describe a new variant of the HBA1 gene co-inherited with a previously described variant of the HBD gene in a diabetic patient tested for HbA1c. Methods: HbA1c and Hb fraction separation were performed using capillary electrophoresis (CE), and the Hb components were confirmed by high-performance liquid chromatography (HPLC). Molecular analysis was performed using next-generation sequencing (NGS). Results: The results of capillary electrophoresis and HPLC tests showed the presence of two anomalous peaks. The molecular study revealed the presence of a new variant of the HBA1 gene, which was named Hb Romagna, and a co-inherited variant of the HBD gene already known as Hb A2-Lampang. Discussion: The coexistence of the two variants poses certain diagnostic challenges with clinical implications that could be relevant to patient management.
Full article
(This article belongs to the Section Conventional Treatment of Thalassemia)
►▼
Show Figures

Figure 1
Open AccessCase Report
Molecular Identification and Familial Segregation of the Hb Malay (HBB:c.59A>G) Variant in a Three-Generation Indonesian Family
by
Chris Adhiyanto, Achmad Zaki, Gema Puspa Sari, Mella Ferania, Yona Mimanda, Laifa Annisa Hendarmin, Suryani, Rini Puspitaningrum, Ayu Latifah, Sakorn Pornprasert, Phakhwan Sampaoloi and Saruda Intachote
Thalass. Rep. 2026, 16(3), 15; https://doi.org/10.3390/thalassrep16030015 - 14 Jul 2026
Abstract
►▼
Show Figures
Background/Objectives: Beta-thalassemia (β-thal) is an inherited hemoglobin disorder caused by mutations in the HBB gene. Hb Malay (HBB:c.59A>G [NM_000518.5], p.Asn20Ser, CD19), a missense substitution (AAC→AGC; Asn→Ser) in exon 1 of the β-globin gene causing β+-thalassemia, has been mainly described in Southeast
[...] Read more.
Background/Objectives: Beta-thalassemia (β-thal) is an inherited hemoglobin disorder caused by mutations in the HBB gene. Hb Malay (HBB:c.59A>G [NM_000518.5], p.Asn20Ser, CD19), a missense substitution (AAC→AGC; Asn→Ser) in exon 1 of the β-globin gene causing β+-thalassemia, has been mainly described in Southeast Asian populations but remains underreported in published Indonesian molecular epidemiology literature. This study aimed to identify the Hb Malay variant and investigate its familial segregation in a large Indonesian extended family. Methods: Cascade molecular screening was initiated following detection of the mutation in a proband referred for genetic testing. Sixteen individuals (one proband and 15 family members) consented to participate. Genomic DNA was extracted from peripheral blood and a 1.9-kb β-globin gene fragment was amplified by PCR. Mutation screening was performed using the Thalassemia GenoArray Kit HBGA-THAL-b31 (Hybribio) and confirmed by Sanger sequencing. A three-generation pedigree was constructed from molecular results and family interviews. Results: The Hb Malay mutation (HBB:c.59A>G) was identified in 13 of 16 individuals: 12 heterozygous carriers (A/G) and one homozygous individual (G/G, subject III-1, aged 15 years). Three individuals showed the normal genotype (A/A). Pedigree analysis demonstrated autosomal recessive inheritance across three generations. No α-globin deletions covered by the GenoArray panel were detected. Conclusions: To our knowledge, this is one of the few documented reports of cascade molecular screening for the Hb Malay (HBB:c.59A>G) variant in a large extended Indonesian family, confirmed by dual molecular methods. These findings illustrate the potential value of cascade molecular screening for early carrier identification in settings where hemoglobin fractionation testing is unavailable, and highlight the need for population-based epidemiological studies to guide thalassemia prevention policy in Indonesia.
Full article

Figure 1
Open AccessArticle
Molecular Characterization of HBB Gene Variations in Beta-Thalassemia Patients from Khyber Pakhtunkhwa, Pakistan
by
Shahzad Ahmad, Laiba Khan, Muhammad Mustafa, Yousaf Khan, Syed Farooq Shah, Fuzail Ahmad, Taimoor Khan, Muhammad Asif Zeb, Qaiser Zaman and Musharraf Jelani
Thalass. Rep. 2026, 16(3), 14; https://doi.org/10.3390/thalassrep16030014 - 10 Jul 2026
Abstract
Background: Beta-thalassemia is a hereditary hematological illness in which beta-globin chain synthesis is missing or decreased, resulting in inefficient erythropoiesis, persistent hemolysis, and anemia. It is most frequently observed in populations with a high rate of consanguineous marriages; affordability becomes a limiting factor
[...] Read more.
Background: Beta-thalassemia is a hereditary hematological illness in which beta-globin chain synthesis is missing or decreased, resulting in inefficient erythropoiesis, persistent hemolysis, and anemia. It is most frequently observed in populations with a high rate of consanguineous marriages; affordability becomes a limiting factor for families with low income. Methods: In this study, we optimized Sanger sequencing of the whole HBB gene covering all three coding exons, two introns, and 5′ and 3′ UTRs. Eleven families were enrolled from Fatimid Foundation Peshawar, Khyber Pakhtunkhwa, Pakistan, with a total of 16 transfusion-dependent patients. An index patient in each family was Sanger sequenced for the HBB gene (n = 11). The variants were classified as per ACMG 2015 guidelines. Results: Sequencing analysis revealed five homozygous pathogenic variations, including three frameshifts: c.27_28dupG (p.Ser10Valfs*14), c.17_18delCT (p.Pro6Argfs17), c.126_129delCTTT (p.Phe42Leufs*19), and two splice-sites (c.92+1G>A, and c.92+5G>C) in nine families. However, two families remained unresolved. The mutation, c.27_28dupG (p.Ser10Valfs*14), was observed in several pedigrees, indicating a probable founder effect or high allele frequency within the affected population. Conclusions: The majority of detected variations in this study were classified as disease-causing, which reside in the first two coding exons and intron-1 of the HBB gene, highlighting its functional importance in the Khyber Pakhtunkhwa population. We recommend performing Sanger sequencing of exon-1-intron-1-exon-2 of the HBB gene with the optimized primers of this study.
Full article
(This article belongs to the Special Issue Genetic Approach in Diagnosis and Following Up of Patients With Thalassemia)
►▼
Show Figures

Figure 1
Open AccessReview
β-Thalassemia in Vietnam: Epidemiology, Molecular Characteristics, Diagnosis, Treatment, and Prevention Strategies
by
Hong-Quan Duong, Thi-Khanh Tran, Thi-Hue Nguyen and Minh-Cong Hoang
Thalass. Rep. 2026, 16(3), 13; https://doi.org/10.3390/thalassrep16030013 - 2 Jul 2026
Abstract
β-thalassemia is one of the most prevalent inherited hemoglobin disorders worldwide and represents a major public health challenge in Southeast Asia, particularly in Vietnam. The disorder is caused by pathogenic variants in the β-globin (HBB) gene, resulting in reduced or absent
[...] Read more.
β-thalassemia is one of the most prevalent inherited hemoglobin disorders worldwide and represents a major public health challenge in Southeast Asia, particularly in Vietnam. The disorder is caused by pathogenic variants in the β-globin (HBB) gene, resulting in reduced or absent β-globin synthesis, ineffective erythropoiesis, chronic hemolytic anemia, and progressive multi-organ complications. Vietnam exhibits a high burden of thalassemia and related hemoglobinopathies, with approximately 13.8% of the population carrying a hemoglobinopathy-associated variant and substantial heterogeneity in distribution across ethnic groups and geographic regions. This review provides a comprehensive overview of the epidemiology, molecular characteristics, diagnostic strategies, treatment approaches, and prevention programs for β-thalassemia in Vietnam. Current evidence indicates that a limited number of recurrent HBB variants account for the majority of β-thalassemia alleles in the Vietnamese population, including codon 17 (A>T) (HBB: c.52A>T), codons 41/42 deletion (-TTCT) (HBB: c.126_129delTTCT), codons 71/72 (+A) (HBB: c.216_217insA), codons 95 (+A) (HBB: c.287_288insA), IVS-I-1 (G>T) (HBB: c.92+1G>T), IVS-I-5 (G>C) (HBB: c.92+5G>C), IVS-II-654 (G>T) (HBB: c.316+197C>T), -28 (A>G) (HBB: c.-78A>G), and -88 (C>T) (HBB: c.-138C>T). Diagnostic strategies generally follow a stepwise approach integrating hematological screening and hemoglobin analysis with confirmatory molecular testing. Advances in molecular diagnostics, particularly targeted polymerase chain reaction (PCR)-based assays and next-generation sequencing (NGS), have significantly improved detection of both carriers and affected individuals. Despite these advances, β-thalassemia continues to impose a considerable clinical, economic, and societal burden because many patients require lifelong blood transfusions, iron chelation therapy, and multidisciplinary management of disease-related complications. Major challenges include limited access to screening, prenatal diagnosis, and genetic counseling services, particularly in rural and ethnic minority populations, as well as the financial and technical barriers associated with advanced molecular diagnostics and emerging therapies. Strengthening nationwide screening programs, expanding access to prenatal and preconception genetic services, improving public awareness, and enhancing healthcare infrastructure are essential for reducing disease incidence and improving patient outcomes. In parallel, the integration of advanced molecular diagnostics and emerging gene-based therapies will be critical for advancing long-term disease control. This review highlights current knowledge gaps and proposes strategic priorities to support evidence-based policy development and comprehensive β-thalassemia prevention and management in Vietnam.
Full article
(This article belongs to the Collection Feature Papers in Thalassemia Reports)
►▼
Show Figures

Figure 1
Open AccessArticle
Comparative Diagnostic Value of 3D Volumetry and Speckle-Tracking over Conventional 2D Echocardiography in the Evaluation of Left Ventricular Function in Pediatric Transfusion-Dependent Beta-Thalassemia
by
Omar Raafat, Ahmed Salama Abouhay, Yasmine El Chazli, Yasser Wali and Hani Mahmoud Adel
Thalass. Rep. 2026, 16(2), 12; https://doi.org/10.3390/thalassrep16020012 - 19 Jun 2026
Abstract
►▼
Show Figures
Background: Left ventricular (LV) dysfunction remains the leading cause of mortality in transfusion-dependent beta-thalassemia (TDßT), yet conventional echocardiography often fails to detect early myocardial impairment. This study aimed to comprehensively evaluate LV function in children with TDßT using three-dimensional echocardiography (3DE) and speckle-tracking
[...] Read more.
Background: Left ventricular (LV) dysfunction remains the leading cause of mortality in transfusion-dependent beta-thalassemia (TDßT), yet conventional echocardiography often fails to detect early myocardial impairment. This study aimed to comprehensively evaluate LV function in children with TDßT using three-dimensional echocardiography (3DE) and speckle-tracking strain analysis, comparing diagnostic performance with conventional two-dimensional (2D) parameters. Results: 50 TDßT patients were compared to 50 matched controls and exhibited preserved conventional LV ejection fraction (EF) on 2D (65.31 ± 7.12% vs. 69.21 ± 3.87%, p = 0.001), but 3DE revealed significant ventricular dilation with higher end-diastolic volume index (75.50 ± 17.99 vs. 65.63 ± 11.86 mL/m2, p = 0.002) and end-systolic volume index (22.28 ± 7.85 vs. 18.21 ± 5.14 mL/m2, p = 0.003). Despite preserved 3D EF (70.79 ± 5.98% vs. 72.07 ± 5.76%, p = 0.276), global longitudinal strain (GLS) was significantly impaired (−18.56 ± 2.37% vs. −21.47 ± 1.86%, p < 0.001). 3D volumetric parameters demonstrated superior diagnostic performance (AUC for LVEDVI Z-score = 0.874) compared to conventional indices. Transfusion duration correlated strongly with ventricular volumes (r = 0.569 for EDV, p < 0.001), while serum ferritin showed no significant correlation with cardiac parameters. Conclusions: Children with TDßT develop early subclinical LV dysfunction detectable by 3DE and strain analysis despite preserved conventional systolic indices. 3D volumetry and GLS should be integrated into routine cardiac surveillance protocols to enable timely therapeutic intervention.
Full article

Figure 1
Open AccessArticle
Transfusion-Transmitted Risk: High Seroprevalence and Genotypic Diversity of Toxoplasma gondii in Thalassemia Major Calls for Donor Screening Policy Review
by
Mitra Sadeghi, Mohammad Naderi Sorki, Bahareh Basirpour, Hossein Karami, Davood Anvari, Seyed Ali Shariatzadeh, Alireza Mohsennejad, Shahabeddin Sarvi, Shirzad Gholami, Seyed Abdollah Hosseini, Sara Gholami, Ahmad Daryani and Sargis. A. Aghayan
Thalass. Rep. 2026, 16(2), 11; https://doi.org/10.3390/thalassrep16020011 - 16 Jun 2026
Abstract
►▼
Show Figures
Objectives: Patients with thalassemia major are at high risk of developing blood-borne infections, including toxoplasmosis, due to their dependence on frequent blood transfusions and underlying immune system disorders. This study was designed to investigate this hidden risk and provide data for policymaking
[...] Read more.
Objectives: Patients with thalassemia major are at high risk of developing blood-borne infections, including toxoplasmosis, due to their dependence on frequent blood transfusions and underlying immune system disorders. This study was designed to investigate this hidden risk and provide data for policymaking in blood transfusion services in a region with a high endemicity. Methods: A total of 300 blood samples from thalassemia patients in northern Iran were collected. Serological testing was conducted to detect IgG and IgM antibodies. DNA extraction followed, with molecular screening performed via PCR. Finally, genotyping of T. gondii was carried out using nested PCR focused on the GRA6 gene. Results: The serological analysis revealed 59.7% of patients exhibited IgG against T. gondii, while only 0.6% tested positive for IgM. The results of the molecular screening revealed 2.7% of patients had DNA of T. gondii. The results of genetic analysis showed 75% had type II, 12.5% had type I, and 12.5% belonged to type III. Conclusions: This study provides serological and molecular evidence of a high chronic Toxoplasma gondii burden in thalassemia patients from northern Iran, an endemic region. A significant association between blood transfusion history and seropositivity, along with parasite DNA detection, suggests elevated exposure risk, though direct transfusion transmission remains unproven. Finding’s support integrating nested PCR with routine serology for diagnosing infection in this population.
Full article

Graphical abstract
Open AccessSystematic Review
Frequency of Thalassemias in the Brazilian Population and Comparison Between Diagnostic Methods: A Systematic Review
by
Eliana A. Santos, Luciana S. Wermelinger and Renato S. Carvalho
Thalass. Rep. 2026, 16(2), 10; https://doi.org/10.3390/thalassrep16020010 - 5 Jun 2026
Abstract
►▼
Show Figures
Thalassemia is an inherited hemoglobin disorder caused by reduced or absent globin chain synthesis, with heterogeneous distribution worldwide and in Brazil. Back-ground/Objectives: This systematic review aimed to estimate the frequency of thalas-semia in the Brazilian population according to thalassemia type, geographic region, and
[...] Read more.
Thalassemia is an inherited hemoglobin disorder caused by reduced or absent globin chain synthesis, with heterogeneous distribution worldwide and in Brazil. Back-ground/Objectives: This systematic review aimed to estimate the frequency of thalas-semia in the Brazilian population according to thalassemia type, geographic region, and population characteristics, as well as to evaluate the impact of diagnostic methods on frequency estimates. Methods: A systematic review was performed following PRISMA 2020 recommendations, in January 2026, including original studies conducted in Brazil-ian populations that reported thalassemia frequency data. Results: Thirty-six studies met the inclusion criteria, of which 77.8% were classified as high quality. The overall average frequency of thalassemia in Brazil was 7.5%, varying according to thalassemia type and diagnostic approach. The mean frequency of alpha thalassemia carriers was 12.3% (range: 5.5–54.0%), with regional variation from 5.79% in the Midwest to 17.3% in the Southeast. The −α3.7 kb deletion was the most frequently reported mutation na-tionwide. Beta thalassemia showed a mean frequency of 2.81% (range: 0.24–18.1%), with regional values ranging from 0.59% in the Southeast to 12.2% in the North and a wide spectrum of pathogenic variants. Distinct frequency patterns were observed in populations with inherent interpretative bias, including individuals with sickle cell trait, systemic lupus erythematosus, microcytosis, and Black populations. Molecular diagnostic methods demonstrated higher sensitivity, enabling the detection of asymp-tomatic carriers and reducing false-negative results. Conclusions: These findings pro-vide a comprehensive epidemiological overview of thalassemia in Brazil and reinforce the importance of molecular diagnostics for accurate screening, genetic counseling, and the development of public health strategies.
Full article

Figure 1
Open AccessArticle
Parental Perspectives on Prenatal Diagnosis and Termination of Pregnancy in Families Affected by β-Thalassemia in Pakistan: A Qualitative Study
by
Iqra Javaid, Muhammad Ameeq and Muhammad Muneeb Hassan
Thalass. Rep. 2026, 16(2), 9; https://doi.org/10.3390/thalassrep16020009 - 9 May 2026
Abstract
Background: Advances in medical genetics and prenatal diagnosis have improved the detection of fetal abnormalities during pregnancy. The findings may lead some couples to consider termination of pregnancy (TOP). This study aimed to explore parental perspectives on prenatal diagnosis and termination of pregnancy
[...] Read more.
Background: Advances in medical genetics and prenatal diagnosis have improved the detection of fetal abnormalities during pregnancy. The findings may lead some couples to consider termination of pregnancy (TOP). This study aimed to explore parental perspectives on prenatal diagnosis and termination of pregnancy among families in which both parents were β-thalassemia carriers and had at least one previously affected child. Methods: A qualitative study was conducted using semi-structured interviews with 30 participants (15 fathers and 15 mothers) recruited from Bahawal Victoria Hospital, Bahawalpur, Pakistan, between November 2024 and February 2025. Eligible couples were registered for chorionic villus sampling (CVS)-based prenatal diagnosis; both parents had confirmed β-thalassemia carrier status, and each family had at least one previously affected child with β-thalassemia major or intermedia. Interview data were analyzed using thematic analysis. Results: Religious beliefs, financial burden, prior experience with affected children, and partner support emerged as major influences on reproductive decision-making. Many parents viewed prenatal diagnosis as important for preparation and informed decision-making. Mothers more often described emotional conflict, stress, and reliance on support, whereas some fathers expressed greater acceptance of termination in the context of severe disease burden. Conclusions: Religious beliefs, prior disease experience, family dynamics, and socioeconomic pressures were important and interrelated influences on decisions about prenatal diagnosis and termination within this study population. Our findings underscore the importance of culturally sensitive, non-directive genetic counseling in low-resource settings. The study was limited by its small sample, single-center design, the use of joint spousal interviews, and the possibility that pre-interview counseling influenced participants’ responses.
Full article
(This article belongs to the Section Conventional Treatment of Thalassemia)
►▼
Show Figures

Figure 1
Open AccessArticle
Association Between Plasma Cystatin C Concentration and Urine Osmolality in Adults with Different Forms of Beta-Thalassemia: A Cross-Sectional Study in Vietnam
by
Loan Do Thi Thanh, Anh Nguyen Ngoc, Kien Nguyen Trung, Ha Nguyen Thi Thu, Dung Nguyen Huu and Thang Le Viet
Thalass. Rep. 2026, 16(2), 8; https://doi.org/10.3390/thalassrep16020008 - 6 May 2026
Abstract
►▼
Show Figures
Objective: To determine plasma cystatin C concentrations, urine osmolality and their relationship with disease severity in beta-thalassemia patients. Methods: A cross-sectional study was conducted on 234 patients with beta-thalassemia, including equal numbers (78 each) of beta-thalassemia major, intermedia, and minor cases, along with
[...] Read more.
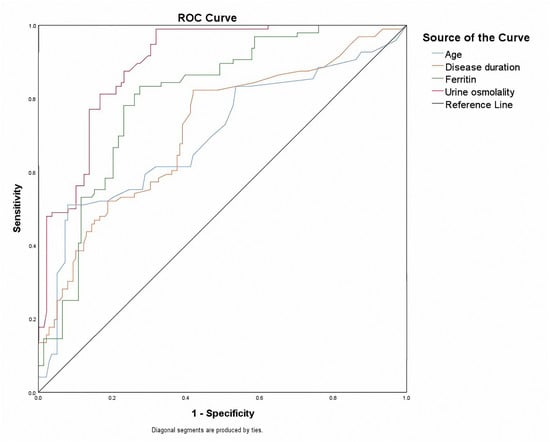
Objective: To determine plasma cystatin C concentrations, urine osmolality and their relationship with disease severity in beta-thalassemia patients. Methods: A cross-sectional study was conducted on 234 patients with beta-thalassemia, including equal numbers (78 each) of beta-thalassemia major, intermedia, and minor cases, along with 78 healthy individuals matched for age, sex, and body mass index, who served as the control group. Plasma cystatin C concentrations were quantified in all subjects using the ELISA method, and urine osmolality level was measured automatically on a FISKE 210 machine (USA). Results: The proportion of beta-thalassemia patients with increased plasma cystatin C concentration was 39.3% and the proportion with a decreased urine osmolality level was 67.5% compared with the control group. Plasma ferritin had predictive value for increased plasma cystatin C concentration (cut-off point: 567.5 ng/mL; AUC = 0.803) and decreased urine osmolality level (cut-off point: 488.15 ng/mL; AUC = 0.820), p < 0.001. Plasma cystatin C concentration increased gradually and urine osmolality level decreased gradually from minor beta-thalassemia to intermedia beta-thalassemia to major beta-thalassemia patients, with p < 0.001. Conclusions: Increased plasma cystatin C concentrations and decreased urine osmolality levels are common and are associated with severity in beta-thalassemia patients.
Full article

Figure 1
Open AccessReview
Recent Advances in Thalassemia Management: From Curative Therapies to Artificial Intelligence
by
Mohamed Medhat Abdelwahab Gamaleldin, Shaimaa Mahmoud Nashat Sayed Abdelhalim and Ivo Abraham
Thalass. Rep. 2026, 16(2), 7; https://doi.org/10.3390/thalassrep16020007 - 22 Apr 2026
Cited by 1
Abstract
Thalassemia is an inherited hemoglobin disorder characterized by chronic hemolytic anemia and substantial long-term healthcare needs. In β-thalassemia major, patients typically require regular red blood cell transfusions with iron chelation to prevent transfusional iron overload. Although supportive care has markedly improved survival, it
[...] Read more.
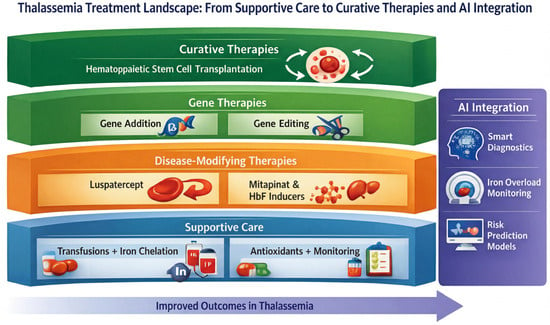
Thalassemia is an inherited hemoglobin disorder characterized by chronic hemolytic anemia and substantial long-term healthcare needs. In β-thalassemia major, patients typically require regular red blood cell transfusions with iron chelation to prevent transfusional iron overload. Although supportive care has markedly improved survival, it is associated with a high treatment burden and does not provide a cure. In recent years, curative and disease-modifying therapies have expanded the treatment landscape. Allogeneic hematopoietic stem cell transplantation (HSCT) offers a potentially curative option for selected patients, while autologous gene therapy and gene-editing approaches have shown the capacity to achieve transfusion independence in clinical studies. In parallel, pharmacologic advances—including luspatercept, a transforming growth factor-beta (TGF-β) ligand trap—have been shown to enhance erythropoiesis and reduce transfusion requirements, and emerging agents such as fetal hemoglobin inducers (e.g., thalidomide) and the oral pyruvate kinase activator mitapivat have demonstrated clinically meaningful hemoglobin improvements in selected populations. Adjunctive strategies, including antioxidants, are under investigation to mitigate oxidative stress, and applications of artificial intelligence are increasingly used to support screening, diagnosis, and longitudinal monitoring of iron overload. This review synthesizes recent advances in curative therapies, novel pharmacologic agents, supportive strategies, and AI-enabled tools and highlights priorities for future clinical development and implementation.
Full article
(This article belongs to the Collection Feature Papers in Thalassemia Reports)
►▼
Show Figures

Figure 1
Open AccessArticle
Implementing Genetic Counseling for Rare Diseases in LMICs: Pediatric and Prenatal Perspectives from India
by
Siddannagoud, Dolat Singh Shekhawat, Kuldeep Singh, Varuna Vyas, Pratibha Singh, Charu Sharma, Navdeep Kaur Ghuman and Tanu Gupta
Thalass. Rep. 2026, 16(2), 6; https://doi.org/10.3390/thalassrep16020006 - 1 Apr 2026
Abstract
Background: This study investigated the clinical characteristics of consultands and examined their perceived personal control, satisfaction, and decision-making regarding genetic testing, as well as the factors influencing these outcomes, at a tertiary care center in northwestern India. Methods: Detailed clinical and family histories
[...] Read more.
Background: This study investigated the clinical characteristics of consultands and examined their perceived personal control, satisfaction, and decision-making regarding genetic testing, as well as the factors influencing these outcomes, at a tertiary care center in northwestern India. Methods: Detailed clinical and family histories were recorded, and trained genetic professionals provided genetic counseling. Perceived personal control (PPC) was assessed pre- and post-counseling using the PPC (nine-item) questionnaire, while post-counseling satisfaction was measured using the six-item Genetic Counseling Satisfaction Scale (GCSS). Outcomes included awareness of genetic disorders, uptake of genetic testing, and reproductive decision-making. Results: A total of 225 consultands (125 pediatric and 100 antenatal) were enrolled. The most common systemic disorders were: inborn errors of metabolism (21.3%), congenital anomalies (15.2%), neurological disorders (15%), primary immunodeficiencies (12.3%), renal genetic disorders (12.2%), respiratory disorders (12%), thalassemia (9%), endocrine disorders (3.9%), and cardiovascular anomalies (3%). In the pediatric group, socioeconomic status (p = 0.048) and higher education levels (p = 0.02) were significantly associated with higher perceptions of adequate counseling time and overall GCSS. None of the examined factors in the prenatal group showed a statistically significant association with satisfaction scores. Consultands primarily concerned with preventing recurrence in future pregnancies showed significantly higher PPC scores both before (p = 0.026) and after counseling (p = 0.009), with the greatest overall improvement in satisfaction (p = 0.044). In the pediatric group, those with an affected family member showed the greatest post-counseling improvement in PPC. Conclusions: Low education, limited awareness, socioeconomic constraints, delayed presentation and low referral rates were key barriers to effective genetic counseling. Addressing these factors can improve consultand awareness, satisfaction, decision-making, and uptake of genetic testing, thereby enhancing reproductive outcomes in high-risk families.
Full article
(This article belongs to the Section Quality of Life)
►▼
Show Figures

Figure 1

Open AccessArticle
Iron Overload and Endocrine Dysfunction in Adults with Transfusion-Dependent Beta-Thalassemia and Growth Retardation: A Correlational Study
by
Muhammad Hammad, Sadaf Fardoos, Khadija Shakoor and Ali Nasir
Thalass. Rep. 2026, 16(1), 5; https://doi.org/10.3390/thalassrep16010005 - 11 Mar 2026
Cited by 2
Abstract
Background and Objective: Iron overload remains a significant clinical concern in patients with transfusion-dependent beta-thalassemia (TDT). This study aims to characterize the iron load and endocrine profile of adult transfusion-dependent beta-thalassemia patients and to evaluate their correlation with growth retardation. Methods:
[...] Read more.
Background and Objective: Iron overload remains a significant clinical concern in patients with transfusion-dependent beta-thalassemia (TDT). This study aims to characterize the iron load and endocrine profile of adult transfusion-dependent beta-thalassemia patients and to evaluate their correlation with growth retardation. Methods: A cross-sectional study was conducted at PIMS Hospital, Islamabad, involving 62 adult patients with homozygous or HbE beta-thalassemia receiving regular blood transfusions. Iron overload was assessed using serum ferritin (SF) and transferrin saturation (TS), while endocrine function was evaluated through measurements of thyroid-stimulating hormone-sensitive (TSH), free thyroxine (FT4), and insulin-like growth factor-1 (IGF-1). Data was analyzed using SPSS v26.0 and R v4.3.1, which included Pearson correlation, chi-square testing, and multivariable regression to explore associations between iron indices and endocrine dysfunction. Results: Serum ferritin demonstrated significant negative correlations with FT4 (r = −0.348, p = 0.005) and IGF-1 (r = −0.302, p = 0.015). MRI T2* pancreas values correlated positively with FT4 (r = 0.268, p = 0.037) and IGF-1 (r = 0.312, p = 0.015). Patients with ferritin > 5000 ng/mL exhibited a higher prevalence of low IGF-1 levels (89.2% vs. 64.0%, p = 0.018). No significant gender-based differences were observed in endocrine parameters. Conclusion: Pancreatic iron burden and elevated serum ferritin were significantly associated with impaired thyroid and growth axis function, highlighting the value of integrating MRI T2* and biochemical markers for early endocrine risk stratification in adult TDT patients.
Full article
Open AccessBrief Report
Hemoglobinopathy Prevention Program in Immigrants: Equality Plus Education Program
by
Duran Canatan, Vincenzo De Sanctis, Joan Lluis Vives Corrons, Giorgio Piacentini, Fatih Kara, Basak Tezel, Aslıhan Ugur Külekci, Özlem Zümrüt, Zekiye Özdemir, Kemal Gürsoy, Gamze Kaymak, Şirin Aydın, Tanju Altunsu, İlhan Aydın, Mustafa Hambolat, Nilgün Keloğlu, Elif Durmaz and Abdullah Solmaz
Thalass. Rep. 2026, 16(1), 4; https://doi.org/10.3390/thalassrep16010004 - 10 Mar 2026
Abstract
►▼
Show Figures
Background and aim: Hemoglobinopathies have become an important public health problem due to global migration. The aim of this project was to address the problem of hemoglobinopathy among immigrants living in Türkiye, Spain, and Italy, in addition to training health managers and Syrian
[...] Read more.
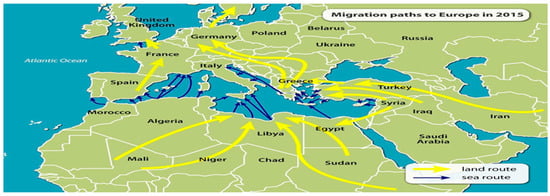
Background and aim: Hemoglobinopathies have become an important public health problem due to global migration. The aim of this project was to address the problem of hemoglobinopathy among immigrants living in Türkiye, Spain, and Italy, in addition to training health managers and Syrian family physicians at immigrant health centers in the southeastern provinces of Türkiye. Material and methods: A three-year international project, named EQUALITY PLUS, was supported by the European Union Erasmus Project. We planned transnational meetings (TPM), vocational education meetings (VET), and Practical Implementation Meetings (PIEM) for the education program. Results: Four TPMs were held in Türkiye, Spain, and Italy, involving a total of 49 professionals. Two VETs were held in Spain and Italy. A total of 23 professionals attended both VETs. Six PIEMs were held in the southern and southeastern Turkish provinces, such as Adana Mersin, Hatay, Gaziantep, Kilis, and Sanliurfa. A total of 442 people, including 373 Syrian family physicians and 69 provincial health managers, were educated in six provinces in Türkiye. Discussion: While the immigrants to Italy and Spain come mainly from Central and North West African maritime routes, immigrants to Türkiye predominantly come from Syria. Among a total of 4 million Syrian immigrants to Türkiye, 200.000 were found to be carriers of thalassemia. In the refugee camps where Syrian immigrants live, the fertility rate is high and the number of sick newborns is increasing, and birth control, genetic counseling, and prenatal diagnosis methods are not sufficient. This project was intended to serve as a guide to prevent hemoglobinopathy in Syrian immigrants. Further projects are needed to address the fertility rate and increased number of sick newborns in these refugee camps. Family physicians at migrant health centers received training on the prevention of hemoglobinopathies. This training included providing detailed genetic counseling to families and providing prenatal diagnosis and preimplantation genetic diagnosis opportunities. Because of the major earthquake that occurred in this region after the project, the work could not continue and preliminary data could not be obtained. Public health services will follow the results of project and the registered number of sick newborns with hemoglobinopathies.
Full article

Figure 1
Open AccessArticle
Cascade Screening of β-Thalassemia in an Indian Family Using Flow Injection Analysis–Triple Quadrupole Mass Spectrometry: Comparison of Micro Sampling Approaches with Conventional Electrophoresis
by
Ankitha K. Puthiyaveettil, Harshini K. Musuvathi and Deepalakshmi D. Putchen
Thalass. Rep. 2026, 16(1), 3; https://doi.org/10.3390/thalassrep16010003 - 24 Feb 2026
Abstract
►▼
Show Figures
Background: β-thalassemia is a rare genetic disorder affecting 1–5% of the global population and poses a health burden due to migration of individuals from endemic regions. Identifying asymptomatic β-thalassemia carriers is essential to prevent the birth of thalassemic babies. A simple, sensitive
[...] Read more.
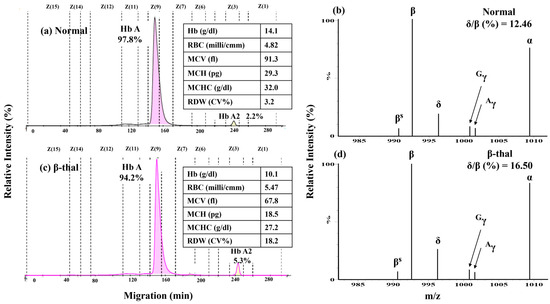
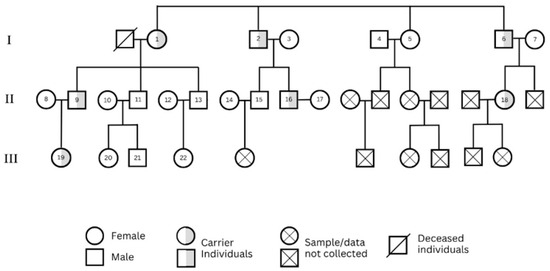
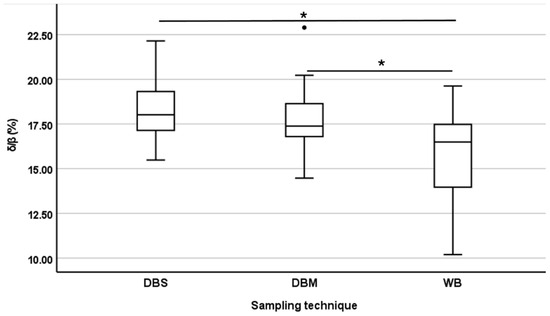
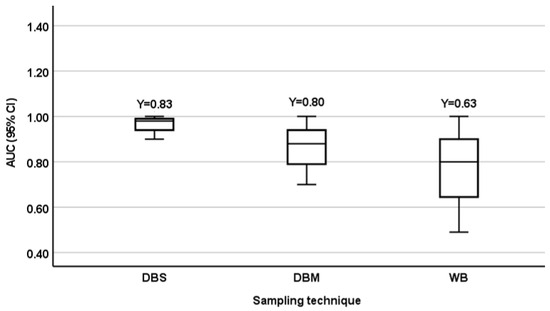
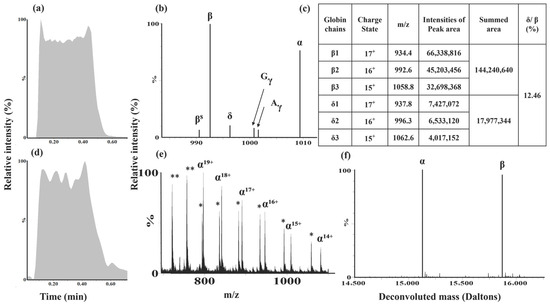
Background: β-thalassemia is a rare genetic disorder affecting 1–5% of the global population and poses a health burden due to migration of individuals from endemic regions. Identifying asymptomatic β-thalassemia carriers is essential to prevent the birth of thalassemic babies. A simple, sensitive method compatible with self-sampling could enhance the detection of β-thalassemia in the population. Methods: Capillary blood was collected via dried blood spot (DBS) and dried blood matrix (DBM) from 18 members (52.9%, 18/34) of a three-generation family. Hemoglobin was extracted, and globin chains were analyzed on a triple quadrupole mass spectrometer (TQMS). δ/β (%) was utilized as a biomarker to identify β-thalassemia. Venous blood collected from positive and negative individuals (n = 11) was further tested to confirm the findings and validated with complete blood count (CBC) and Capillary Electrophoresis (CE). Results: β-thalassemia was detected in seven individuals: three from generation I, three from generation II, and one from generation III. CBC showed thalassemia indices, while CE demonstrated elevated HbA2 consistent with β-thalassemia. Molecular sequencing of two samples confirmed the heterozygous c.92 + 5 G > C mutation in the β-globin gene. The overall prevalence of β-thalassemia in the family was 20.6% (7/34). High clinical performance was achieved across sample types, with 100% sensitivity for DBS, 100% specificity for DBM, and an overall accuracy of 91% when compared with CE. Conclusions: TQMS in combination with CBC parameters successfully identified asymptomatic heterozygous β-thalassemia carriers using self-sampling techniques. Cascade screening within affected families emerges as a possible strategy for early detection of β-thalassemia pending comprehensive validation.
Full article

Figure 1
Highly Accessed Articles
Latest Books
E-Mail Alert
News
14 July 2025
Meet Us Virtually at the 1st International Online Conference on Hematology, 28–30 October 2026
1 September 2026
MDPI INSIGHTS: The CEO’s Letter #38 – 2 Million Published Articles, Outstanding Reviewers, Michele Parrinello Award, AIS 2026 & WSF-12
Topics
Topic in
Hematology Reports, Thalassemia Reports, Hemato, JCM, Cancers
New Drugs and Combinations in the Management of Hematological Malignancies and Rare Diseases
Topic Editors: Claudio Cerchione, Aurelio MaggioDeadline: 30 June 2027
Conferences
Special Issues
Special Issue in
Thalassemia Reports
Advances in Hematopoietic Stem Cell Research and Therapeutic Strategies for Thalassemia
Guest Editors: Sandeep Yadav, Aloukick Kumar SinghDeadline: 20 October 2026
Special Issue in
Thalassemia Reports
Genetic Approach in Diagnosis and Following Up of Patients With Thalassemia
Guest Editor: Duran CanatanDeadline: 31 December 2026
Topical Collections
Topical Collection in
Thalassemia Reports
Feature Papers in Thalassemia Reports
Collection Editors: Aurelio Maggio, Khaled Musallam