


Abstract
Drought during reproductive development and grain filling is a major constraint to bread wheat productivity in rainfed environments. In the present study, we employed genome-wide association analysis in an untapped diversity panel of wheat genotypes relevant to natural dryland conditions. A total of 187 genotypes were evaluated for drought-related physiological, biomass, and architectural traits under terminal rainfed stress in Azerbaijan. Relative water content, plant height, fresh weight, dry weight, flag leaf length, and flag leaf width were assessed at the milk ripening stage. These data were subjected to genome-wide association analysis using 19,737 SNP markers to identify loci and epistatic interactions involved in the determination of these traits. The panel showed broad phenotypic variation and significant genotypic effects for all traits, with broad-sense heritability ranging from 0.991 for plant height to 0.385 for flag leaf width. GWAS identified a major locus for relative water content on chromosome 2D at SNP marker AX-86184518, which explained 11.94% of the genotypic variation. Candidate-gene analysis identified the proximal WEB-family-like gene TraesCS2D03G1001000 as a putative positional candidate. Plant height showed strong additive loci, mainly on chromosomes 2A and 4A, whereas biomass and flag leaf traits showed suggestive additive loci and epistatic interactions. These findings identify candidate loci and putative interaction patterns associated with drought-related traits, which require further validation across independent populations and environments.
1. Introduction
Bread wheat (Triticum aestivum L.) is central to global food security because it contributes substantially to dietary energy, plant-based protein and other nutritional components in human diets [1]. Recent FAOSTAT data indicate that global wheat production remains close to 800 million tonnes annually from more than 200 million hectares, confirming wheat as one of the most widely grown food crops worldwide [2]. Maintaining this production level under changing climatic conditions is becoming increasingly difficult. Heat, drought and compound dry–hot events are expected to occur more frequently during wheat-growing seasons, with particularly strong consequences for rainfed production systems [3,4]. Because a large share of global wheat is produced under rainfed conditions, yield stability is strongly influenced by the timing, duration and intensity of water deficit [5]. Drought occurring during reproductive development and grain filling is especially damaging, as it restricts canopy function, accelerates senescence and reduces assimilate availability for grain formation [6,7].
Drought reduces wheat performance through a sequence of linked physiological and growth responses, beginning with impaired plant water status and progressing to reduced carbon assimilation, canopy development and biomass accumulation. Leaf relative water content (RWC) provides a direct estimate of leaf hydration because it relates fresh tissue mass to fully turgid and oven-dry mass [8]. In bread wheat, recent morpho-physiological screening under well-watered and drought-stressed conditions confirmed that RWC varies significantly among genotypes and declines under water deficit, while its association with excised leaf water retention, relative water loss and membrane stability reflects broader differences in leaf water status and drought response [9]. Water-relation analyses in contrasting wheat genotypes further indicate that drought responsiveness depends on the coordination between leaf water status, stomatal regulation, osmotic adjustment and cell wall elasticity rather than on leaf water content alone [10]. Thus, RWC, fresh weight (FW) and dry weight (DW) provide complementary information on the physiological and growth components of drought response, making them suitable traits for genetic analysis under field moisture limitation.
Plant architecture is another important component of dryland adaptation because it influences lodging resistance, biomass distribution, harvest index and suitability to target production environments. Plant height (PH) is not simply a trait to be reduced: excessively tall plants are more prone to lodging, whereas excessively short plants may show reduced biomass and yield potential [11]. Recent genetic analysis of plant-height-related traits in wheat also showed that dryland cultivars were approximately 10 cm taller than irrigated cultivars, supporting the view that optimal plant architecture may differ between water-limited and irrigated environments [11]. The Green Revolution semi-dwarfing alleles Rht-B1b and Rht-D1b improved lodging resistance and harvest index, but gibberellin-insensitive reduced-height alleles can shorten coleoptiles and reduce early seedling vigor, which may be disadvantageous under dry sowing or deep-sowing conditions [12,13]. Alternative reduced-height loci therefore remain relevant for dryland breeding, particularly when moderate height reduction is needed without compromising early establishment; Rht25, for example, has been characterized as a gibberellin-sensitive dwarfing locus affecting PH in wheat [14]. In addition to plant height, flag leaf length (FLL) and flag leaf width (FLW) are agronomically relevant because the flag leaf is a major photosynthetic source during grain filling. Genetic dissection of flag leaf morphology under contrasting water regimes has shown that these traits are quantitatively inherited and responsive to water availability [15]. Flag-leaf-related traits, including length and width, have also been associated with plant architecture and grain yield in wheat, supporting their value as target traits in genetic analysis [16]. More recently, natural allelic variation has been shown to contribute to diversity in the regulation of wheat flag leaf morphology, reinforcing the value of these traits for association mapping in diverse germplasm [17]. However, the extent to which flag leaf morphology is genetically coordinated with foliar water status and biomass accumulation under rainfed field stress remains insufficiently resolved.
The genetic dissection of drought-related traits is complicated by their quantitative inheritance and sensitivity to environmental conditions. Drought adaptation in wheat is generally controlled by multiple physiological, developmental and genetic mechanisms rather than by single major-effect loci [18]. Biparental quantitative trait locus (QTL) mapping has been useful for detecting segregating loci in specific populations, but its allelic coverage is limited by the two parental genotypes used in each cross. Genome-wide association study (GWAS) provides a complementary approach because it exploits historical recombination and linkage disequilibrium in diverse panels to detect marker–trait associations at higher resolution. In wheat, GWAS has been used to identify loci associated with seedling drought tolerance [19], osmotic adjustment and related drought-adaptive traits in durum wheat [20], and drought-induced proline and hydrogen peroxide accumulation under natural field and rain-out shelter conditions [21]. In Turkish winter wheat germplasm, the integration of GWAS and selective sweep analysis identified genomic regions and candidate genes associated with improved grain yield under drought conditions [22]. The biological interpretation of these associations has been strengthened by the availability of annotated wheat genome resources, including the reference genome and multi-genome assemblies [23,24]. More recent studies have identified single-nucleotide polymorphism (SNP) markers, candidate genes and gene networks associated with drought tolerance and drought-response indices in wheat [25,26,27]. However, most GWAS studies still emphasize additive marker effects, whereas non-additive marker interactions may also contribute to trait variation under field stress. Therefore, combining additive GWAS with epistatic interaction analysis may provide a more complete view of the genetic basis of drought-related variation in adapted wheat germplasm.
Despite increasing progress in wheat drought genetics, important knowledge gaps remain for rainfed field environments. Many studies have used seedling-stage assays, hydroponic or PEG-induced stress, or controlled drought treatments that do not fully reproduce the fluctuating moisture availability, temperature conditions and soil constraints experienced in dryland production systems. In addition, the genetic coordination among foliar hydration, flag-leaf growth-related traits and plant architecture remains insufficiently understood in locally and regionally adapted germplasm evaluated under real field moisture limitation. Moreover, drought-associated loci identified in individual environments may be environment-specific, emphasizing the need for validation across multiple environments and independent populations before their broader application in breeding. Addressing this gap is important because drought adaptation in breeding programs depends not only on statistically significant markers, but also on markers associated with biologically interpretable traits expressed in target production environments.
As an integral geographical part of a center of wheat diversity and origin, Azerbaijan provides a relevant dryland environment for studying rainfed wheat production in a broader global context. Under these natural dryland conditions, wheat production is exposed to variable precipitation and terminal-season moisture limitation, while locally and regionally adapted germplasm may retain allelic variation shaped by recurrent water-limited conditions [28]. Previous studies on Azerbaijani wheat germplasm have revealed substantial variation in drought-responsive physiological, biochemical, ultrastructural and molecular traits [7,29,30,31,32]. However, the genome-wide basis of this adaptive variation remains insufficiently characterized under rainfed field conditions.
The present study evaluated a diverse panel of 187 bread wheat genotypes under rainfed field conditions in Azerbaijan. The objectives were to: (1) characterize phenotypic variation, broad-sense heritability and inter-trait relationships for RWC, PH, FW, DW, FLL and FLW under rainfed field conditions; (2) identify additive marker–trait associations using SNP-based GWAS; (3) evaluate epistatic marker–marker interactions contributing to drought-related trait variation; and (4) identify candidate genes located near significant markers. By linking physiological, architectural and biomass traits with genome-wide marker information, this study aims to clarify the genetic basis of drought-related variation in field-grown wheat and to provide additional insight into both additive and putative non-additive genetic effects underlying these traits.
2. Materials and Methods
2.1. Plant Material
The plant material utilized in the present study consists of a newly established diversity set of 187 genotypes of bread wheat (Triticum aestivum L.) adapted to natural rainfed conditions representing a wide geographic region across the center of wheat diversity and its origin (Supplementary Table S1). This collection was designed to capture a broad range of phenotypic and genetic diversity relevant for drought adaptation under rainfed conditions.
2.2. Experimental Design and Growth Condition
Field experiments were conducted during the 2022/2023 growing season at the Gobustan Regional Experimental Station of the Research Institute of Crop Husbandry, Azerbaijan (800 m a.s.l.) (Supplementary Figure S1). The soil in this region is classified as light chestnut with a slightly alkaline pH. It is characterized by low humus content (1.5–1.8% in the topsoil), low nitrogen and phosphorus availability, and moderate potassium levels. The hydro-meteorological conditions for the 2022/2023 growing season are presented in detail in Supplementary Table S2. Total precipitation during the cropping season was 356.3 mm, which is below the long-term average of 406.0 mm. A critical moisture deficit occurred in June, with only 4.1 mm of rainfall accompanied by relatively high temperatures, causing pronounced drought stress during the reproductive stages of wheat development. The experiment followed a randomized complete block design (RCBD) with three replications per genotype. Each replicate plot measured 1.5 × 1.0 m (1.5 m2) and consisted of seven rows spaced 15 cm apart. Sowing was performed at a density of 450 germinating seeds per m2.
2.3. Phenotypic Evaluation of Drought-Related Traits
Phenotypic measurements were conducted during the first ten days of June 2023, when the panel was generally within the milk-ripening period. To reduce differences in developmental stage among genotypes, the order of phenotyping within this 10-day period was adjusted according to the observed developmental status of the plants: more advanced genotypes were evaluated earlier, whereas less advanced genotypes were evaluated later. This approach was intended to obtain measurements at broadly comparable developmental stages; however, heading date was not included as an additional covariate in the statistical analyses. Six drought-related agronomic and physiological traits were evaluated: PH, FLL, FLW, FW, DW, and RWC. For each genotype, three plants were randomly selected from each of the three replicate plots. Measurements were performed on individual plants, and the mean of the three plants within each plot was used as the corresponding replicate value for statistical analysis (Supplementary Table S9). Leaf sampling was conducted during the morning hours and completed before 12:00 h to minimize potential diurnal variation in leaf water status.
PH was measured from the soil surface to the top of the spike, excluding awns. Fully developed flag leaves from the same sampled plants were used for the assessment of FLL, FLW, FW, DW, and RWC. FLL was measured from the base to the tip of the fully developed flag leaf, while FLW was measured at the widest part of the same leaf using a ruler with a 1 mm scale resolution [15]. Following the morphological measurements, the same flag leaves were used for laboratory determinations. FW was recorded immediately after sampling. The leaves were then immersed in distilled water for 24 h to reach full turgidity, after which turgid weight (TW) was recorded. Subsequently, the same samples were dried at 80 °C until constant weight and weighed to determine DW.
RWC was calculated according to Turner [8] using the following formula:
where FW is the initial fresh weight of the leaf, TW is the turgid weight after rehydration, and DW is the dry weight after oven-drying.
RWC (%) = [(FW − DW)/(TW − DW)] × 100
2.4. Broad Sense Heritability (H2)
Descriptive statistics were calculated using R software v3.6 (R Foundation for Statistical Computing, Vienna, Austria). A two-way analysis of variance (ANOVA) was performed to assess the effects of genotype and replication, and the emmeans package (https://CRAN.R-project.org/package=emmeans, accessed on 11 September 2026) was used for mean comparisons. Variance components for broad-sense heritability were estimated using a linear mixed model implemented with the nlme package (https://CRAN.R-project.org/package=nlme, accessed on 11 September 2026). The mixed model was:
where Yij is the phenotypic value of the i-th genotype in the j-th replication, μ is the overall mean, Gi is the random effect of genotype, Rj is the random effect of replication (block), and εij is the residual error. Genotype was treated as a random effect in the mixed model to estimate genotypic variance for broad-sense heritability.
Yij = μ + Gi + Rj + εij
Broad-sense heritability on a genotype-mean basis was calculated as:
where Vg is genotypic variance and Verr is the error variance and r = 3 is the number of field replications. Because the experiment was conducted at a single location during one growing season, genotype-by-environment (G × E) variance could not be estimated and was therefore not included in the calculation. Accordingly, the reported H2 values represent single-environment estimates.
2.5. Pearson Correlation Analysis
Pearson correlation analysis was performed to evaluate the relationships among RWC, PH, DW, FW, FLL, and FLW. Correlation coefficients were calculated using genotype means averaged across the three field replications. The resulting correlation matrix was visualized as a heatmap to identify coordinated variation among physiological, biomass-related, and plant-architecture traits under rainfed field conditions.
2.6. Genotyping of the Diversity-Set
The wheat accessions were genotyped by TraitGenetics GmbH (Gatersleben, Germany) using the Illumina Infinium XT 25K SNP array (Illumina, Inc., San Diego, CA, USA). An initial set of 24,146 SNPs was subjected to quality control. SNP markers with ≥10% missing genotype calls were excluded, and only polymorphic markers with a minor allele frequency (MAF) > 0.05 were retained. Missing genotype calls in the retained marker set were imputed using SAS v9.4 (SAS Institute Inc., Cary, NC, USA) prior to association analysis. This filtering resulted in a final dataset of 19,737 high-quality polymorphic SNPs used for all downstream analyses. Details of these markers are described in Zakieh et al. [33,34].
2.7. GWAS
Genome-wide association mapping was performed following the GRAMMAR method described by Aulchenko et al. [35]. GRAMMAR was selected as a computationally efficient mixed-model framework for genome-wide marker testing that accounts for background genetic structure and is consistent with analytical approaches previously applied in comparable wheat association studies. A marker-based identity-by-state (IBS) kinship matrix was calculated using the A.mat function of the rrBLUP package (https://CRAN.R-project.org/package=rrBLUP, accessed on 11 September 2026) in R, following Benaouda et al. [36]. GWAS was performed using the GRAMMAR approach implemented in PROC MIXED of SAS v9.4. The first three principal components and the kinship matrix were included as covariates to account for population structure and genetic relatedness. The number of principal components was selected a priori following the population-structure correction strategy described by Benaouda et al. [36], rather than through a dataset-specific scree-plot criterion. Model calibration was evaluated using quantile–quantile (QQ) plots and the genomic inflation factor (λGC) for each trait. For the analysis, the first three principal components were included as co-factors to control for population structure in the analysis additionally. The methods of the analysis were used and described in detail by Reinert et al. [37] and Naz et al. [38]. We used a linear mixed model to calculate the QTLs as presented below:
where Yij is the phenotypic value; μ is the general mean; Mi is the fixed effect of i-th marker genotype; Lj (Mi) is the random effect of j-th cultivar nested within i-th marker genotype and εij is the residual. Significant markers were detected following a correction using the probability of false discovery rate (FDR), implemented in the SAS procedure PROC MULTTEST according to Benjamini and Yekutieli [39]. To account for multiple testing across all SNPs, genome-wide significance was additionally assessed using a Bonferroni correction based on the total number of markers tested (p ≤ 2.53 × 10⁻⁶). To assess the sensitivity of the detected associations to GWAS model choice, an additional kinship-based single-locus mixed linear model (MLM) analysis was performed for all six traits using the same panel of 187 genotypes and the same set of 19,737 SNP markers. The first three principal components were included as fixed covariates, and genomic relatedness was incorporated through an additive kinship matrix. The alternative MLM was used as a sensitivity analysis rather than as the primary discovery model. Cross-model results for the selected marker–trait associations are provided in Supplementary Table S8.
Yij = μ + Mi + Lj (Mi) + εij
To characterize the local genomic context of the major corrected associations, linkage disequilibrium (LD) was evaluated within ±5 Mb of the lead RWC marker AX-86184518 on chromosome 2D and the major PH markers AX-158540611 on chromosome 2A and AX-94514459 on chromosome 4A. Pairwise LD was expressed as squared allele-dose correlation (r2) using available non-missing genotype calls from the 187-genotype panel. Local association plots were generated using physical SNP positions and −log10(P) values, together with pairwise LD heatmaps for the SNPs within each regional interval.
2.8. Epistatic Interaction Analysis
Pairwise epistatic marker interactions were analyzed using a two-way multilocus approach implemented in the PROC MIXED procedure of SAS, following the general modelling and marker pre-screening strategy described by Benaouda et al. [36]. To reduce the computational burden, markers with a single-marker association p < 0.7 were retained for interaction screening. This permissive threshold was used as a pre-screening criterion rather than as a significance threshold, allowing markers with relatively weak marginal effects to remain eligible for interaction testing while substantially reducing the number of pairwise comparisons. The pre-screen retained 14,500, 15,805, 14,458, 14,061, 14,404, and 14,508 markers for RWC, PH, DW, FW, FLL, and FLW, respectively, corresponding to 105,117,750, 124,891,110, 104,509,653, 98,848,830, 103,730,406, and 105,233,778 unique marker-pair comparisons. No marker × environment interaction terms were fitted because the study was conducted in a single field environment.
The following model was used:
where Yijk is the phenotypic value, μ is the general mean, M1i is the fixed effect of the i-th genotype class of the first marker, M2j is the fixed effect of the j-th genotype class of the second marker, M1j × M2j is the fixed interaction effect between the two marker genotype classes, and εijk is the residual error.
Yijk = μ + M1i + M2j + M1i × M2j + εijk
Marker-by-marker interaction effects were assessed using the probability value of the interaction term. To account for the large number of interaction tests, interaction p-values were additionally reassessed using a trait-specific Bonferroni correction based on the total number of unique marker pairs after pre-screening. The resulting significance thresholds were 4.76 × 10⁻¹⁰ for RWC, 4.00 × 10⁻¹⁰ for PH, 4.78 × 10⁻¹⁰ for DW, 5.06 × 10⁻¹⁰ for FW, 4.82 × 10⁻¹⁰ for FLL, and 4.75 × 10⁻¹⁰ for FLW. Only interactions passing the corresponding corrected threshold were considered statistically supported.
Reciprocal duplicate pairs, in which the same marker combination appeared as Marker A × Marker B and Marker B × Marker A, were collapsed into a single unique marker pair. For representative marker pairs, allele-combination means were calculated to describe phenotypic variation among the observed two-locus genotype combinations. The minimum and maximum allele-combination means were identified, and the mean range was calculated as the difference between them. Five representative Bonferroni-supported interactions per trait were selected for detailed presentation, while the reciprocal-collapsed interaction results and corrected significance status are provided in Supplementary Table S4. Detailed allele-combination means and class sizes for the representative interactions are provided in Supplementary Table S5. Because the analysis was conducted in a single field environment, all statistically supported interactions were nevertheless considered putative and require validation in independent populations and additional environments.
2.9. Determination of Genes Colocalizing with QTL
To identify putative candidate genes associated with the significant GWAS loci, lead SNP markers were anchored to the wheat reference genome (IWGSC RefSeq v2.1) based on their physical positions in the Ensembl-plant database (https://plants.ensembl.org/, accessed on 11 September 2026) [40]. Candidate gene mining was performed using the BioMart interface (https://plants.ensembl.org/biomart/, accessed on 11 September 2026) [41]. A predefined ±500 kb physical window around each lead SNP was used as an exploratory interval for positional candidate-gene screening. Because LD was evaluated separately for the major loci, this fixed window was used as a standardized positional screening interval rather than as an interval derived from LD decay. Accordingly, it was not considered an LD-defined causal region. All annotated gene models whose start or end coordinates fell within this interval were retrieved and considered as putative candidate genes. Functional annotation of candidate genes was obtained based on Gene Ontology (GO) terms describing molecular functions and biological processes, using information available in the UniProt database (https://www.uniprot.org/, accessed on 11 September 2026) [42].
2.10. In Silico Expression Analysis of Candidate Genes
To support the functional interpretation of genes located within candidate QTL intervals, publicly available wheat transcriptome data were inspected using the Wheat Expression Browser, which is based on the expVIP platform (https://www.wheat-expression.com/, accessed on 11 September 2026) [43] and the wheat expression atlas described by Ramírez-González et al. [44]. For candidate genes identified in IWGSC RefSeq v2.1, corresponding RefSeq v1.1 gene models were used when available, because the expression browser is primarily organized according to RefSeq v1.1 annotations. Expression profiles were examined across available non-stress and abiotic-stress-related samples. The abiotic-stress-associated datasets included drought, heat, and combined drought–heat treatments of 1-week-old seedling leaves (0, 1, and 6 h), PEG 6000-treated seedlings (2–12 h), phosphate-starved shoots (10 d), cold-treated shoots (2 weeks), and additional abiotic-stress-associated samples including developing spike material. The corresponding condition-level information is summarized in Supplementary Table S7. Transcript abundance was expressed as transcripts per million (TPM), and mean TPM values were compared descriptively between abiotic-stress-associated and non-abiotic sample groups. This analysis was used only as supportive evidence for prioritizing positional candidate genes and was not treated as experimental validation of gene function.
3. Results
3.1. Phenotypic Variation, Heritability and Inter-Trait Correlations
Substantial phenotypic variation was observed among the 187 bread wheat genotypes for all six drought-related traits under rainfed field conditions (Table 1). RWC ranged from 41.14% to 95.97%, with a mean value of 68.15%, indicating broad variation in plant water status under terminal drought. PH varied from 65.00 to 135.33 cm, whereas DW and FW ranged from 0.05 to 0.29 g and from 0.17 to 1.03 g, respectively. FLL ranged from 14.00 to 31.93 cm, while FLW varied from 1.00 to 2.70 cm.
Table 1.
Phenotypic variation and broad-sense heritability of drought-related traits under rainfed field conditions.
The coefficient of variation was highest for the two biomass-related traits, DW (31.39%) and FW (31.02%), suggesting greater relative dispersion for biomass accumulation under rainfed conditions. In contrast, RWC, PH, FLL and FLW showed moderate coefficients of variation, ranging from 13.74% to 14.53%. Genotypic effects were highly significant for all traits (p < 0.001), confirming the presence of substantial genetic variation within the panel.
Broad-sense heritability differed considerably among traits. The highest single-environment estimate was observed for PH (H2 = 0.991), followed by DW (H2 = 0.785), FW (H2 = 0.775), FLL (H2 = 0.654), RWC (H2 = 0.639), and FLW (H2 = 0.385). The high H2 observed for PH reflects strong differentiation among genotypes relative to residual variation within the present field trial. However, because G × E variance could not be estimated, this value should not be interpreted as an across-environment heritability estimate.
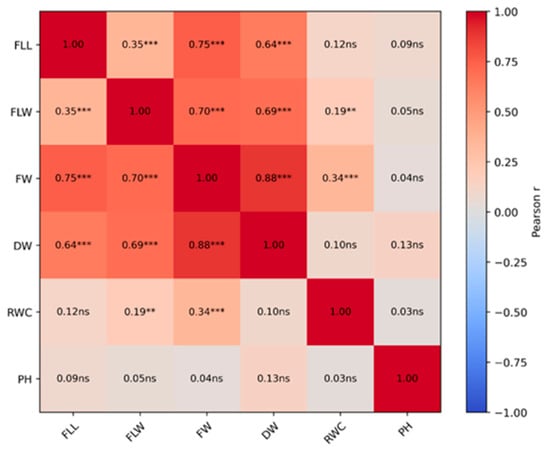
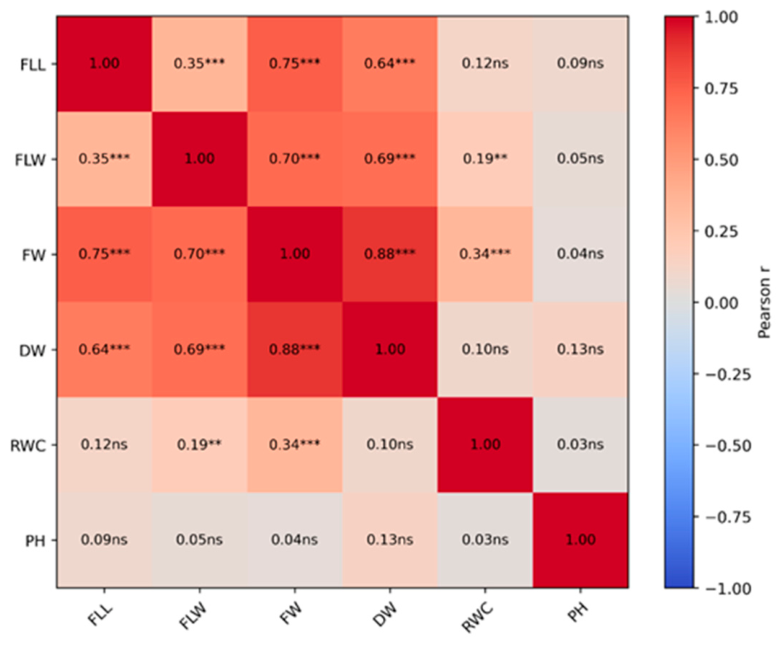
Pearson’s correlation analysis was performed to evaluate the relationships among physiological, biomass-related, and architectural traits (Figure 1). The strongest positive correlation was observed between FW and DW (r = 0.88, p < 0.001), indicating a close relationship between fresh and dry biomass accumulation. FW was also strongly and positively correlated with FLL (r = 0.75, p < 0.001) and FLW (r = 0.70, p < 0.001), while DW showed positive correlations with FLW (r = 0.69, p < 0.001) and FLL (r = 0.64, p < 0.001). A moderate positive correlation was detected between FLL and FLW (r = 0.35, p < 0.001), suggesting partial coordination between the two-flag leaf architectural traits.
Figure 1.
Pearson correlation matrix of drought-related traits in bread wheat under rainfed field conditions. Pearson correlation coefficients (r) were calculated among flag leaf length (FLL), flag leaf width (FLW), fresh weight (FW), dry weight (DW), relative water content (RWC), and plant height (PH). The heatmap shows the strength of pairwise phenotypic correlations among traits; color intensity reflects the magnitude of the correlation coefficient. Asterisks indicate statistical significance: p < 0.01 (**), and p < 0.001 (***), while “ns” indicates non-significant correlations.
RWC showed a moderate positive correlation with FW (r = 0.34, p < 0.001) and a weaker but significant positive correlation with FLW (r = 0.19, p < 0.01). In contrast, RWC was not significantly correlated with FLL (r = 0.12), DW (r = 0.10), or PH (r = 0.03). PH also showed no significant correlations with the other measured traits, including FLL (r = 0.09), FLW (r = 0.05), FW (r = 0.04), DW (r = 0.13), and RWC (r = 0.03). Overall, the correlation structure suggests that biomass accumulation and flag leaf morphology were closely associated, whereas PH behaved largely independently from the other drought-related traits in this panel. The partial association of RWC with FW and FLW further indicates that foliar water status was related to some, but not all, components of biomass accumulation and leaf architecture under rainfed conditions.
3.2. Marker–Trait Associations Using GWAS and Multiple-Testing Correction
Genome-wide association analysis was performed using 19,737 high-quality polymorphic SNP markers. The strength and number of marker–trait associations differed markedly among the six evaluated traits (Supplementary Table S3). At the suggestive threshold of p < 1 × 10⁻⁴, PH showed the largest number of associated SNPs, with 114 SNPs detected. RWC showed ten SNPs with p < 1 × 10⁻⁴, followed by FLW with nine SNPs, DW with four SNPs, FW with two SNPs, and FLL with one SNP.
After multiple-testing correction, PH showed 616 SNPs passing the FDR threshold, of which eight also passed the Bonferroni threshold. For RWC, one SNP passed both the FDR and Bonferroni thresholds. In contrast, no SNPs passed either the FDR or Bonferroni threshold for DW, FW, FLL, or FLW. Therefore, associations detected for these traits were interpreted as preliminary statistically identified marker–trait effects.
Quantile–quantile (QQ) plots were generated for all six traits to compare the observed and expected distributions of association p-values (Supplementary Figure S2). For most traits, the observed p-values followed the expected distribution closely, with deviations mainly in the upper tail, whereas PH showed the strongest departure from the null expectation.
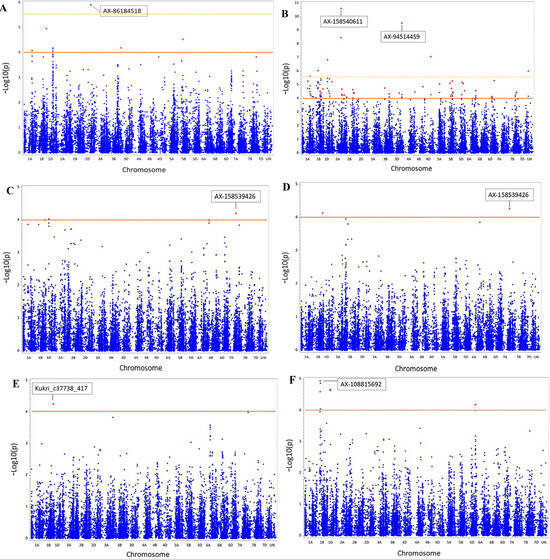
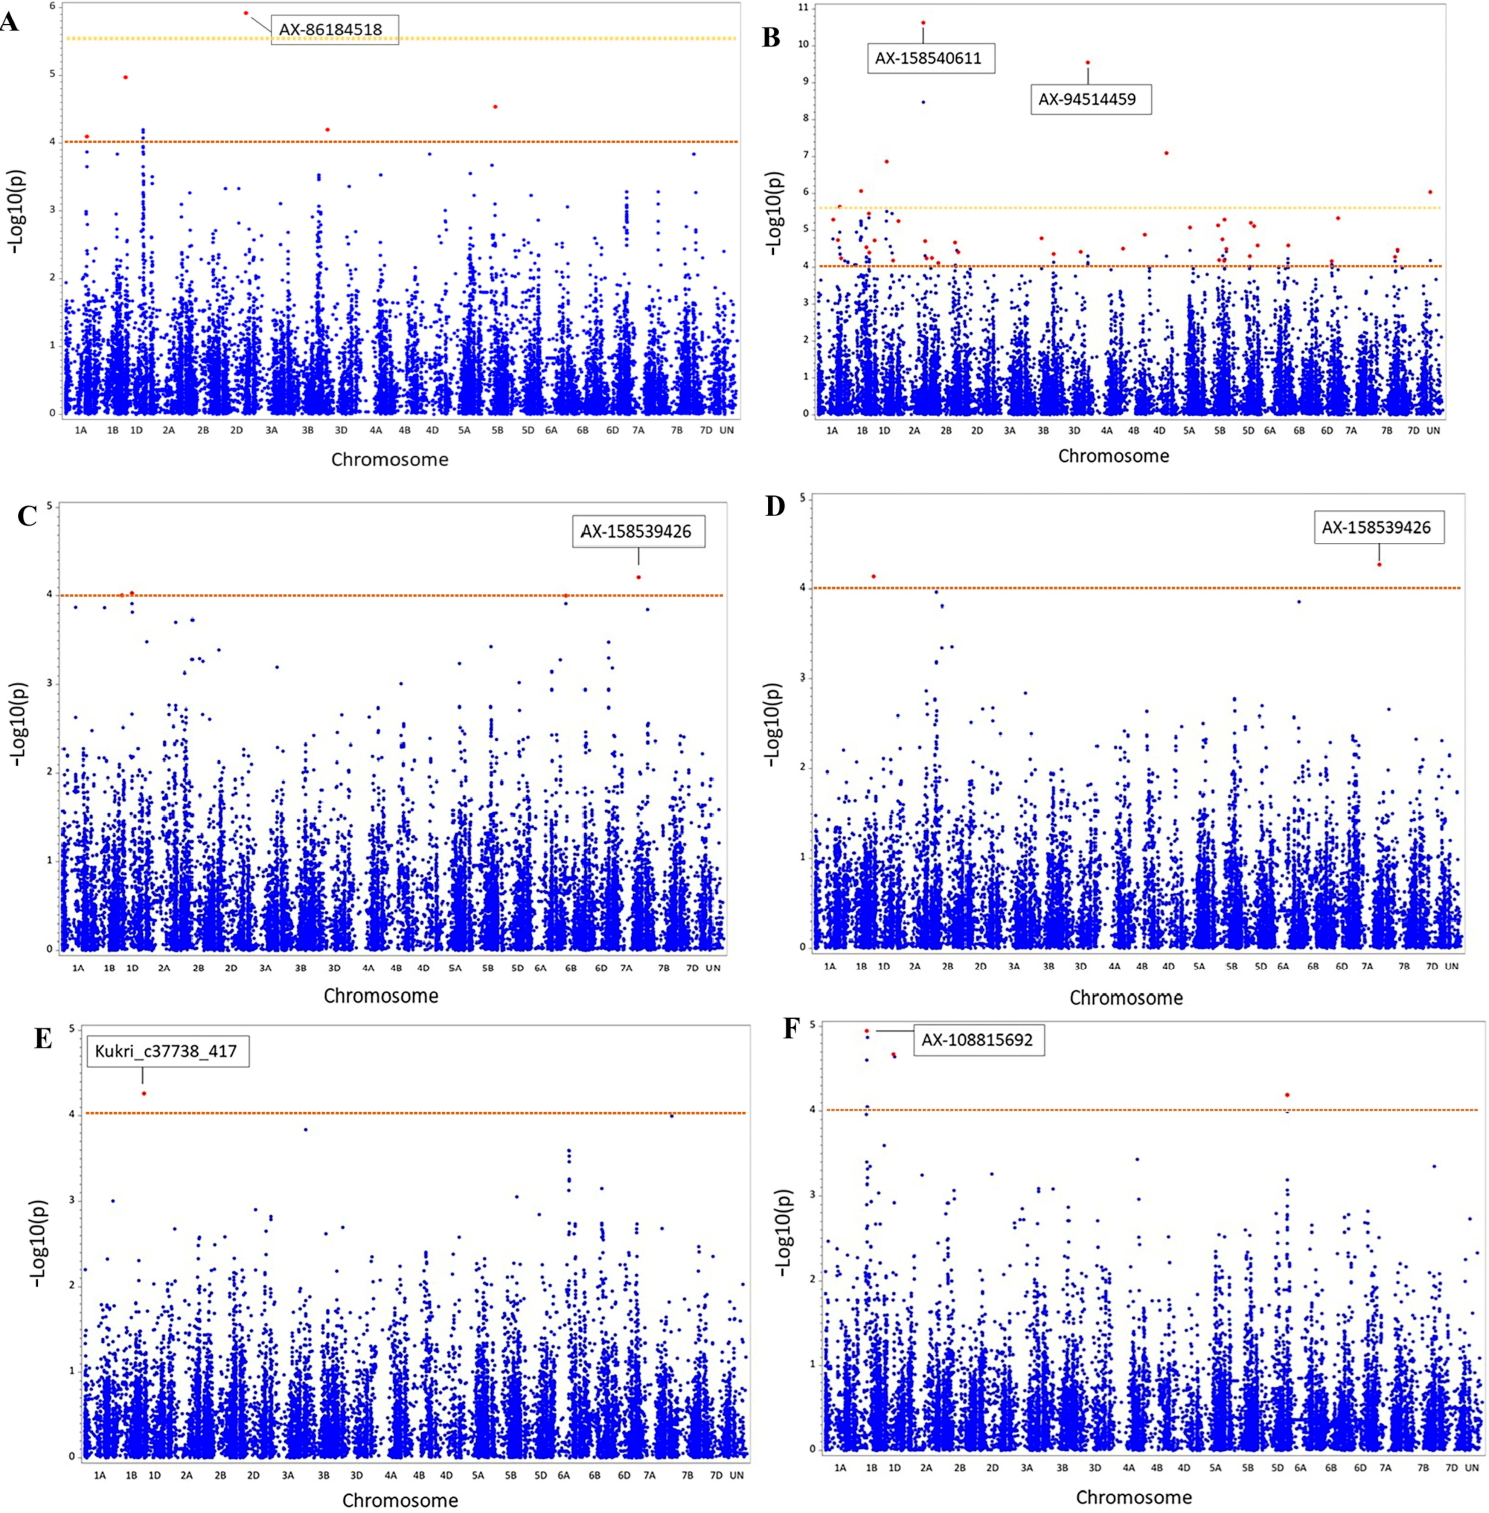
The Manhattan plots illustrated this trait-specific pattern, with the most pronounced associations observed for PH, one corrected association for RWC, and mainly suggestive marker–trait associations for biomass and flag leaf traits (Figure 2). In these plots, red points indicate peak markers retained after marker selection based on association strength (p < 1 × 10⁻⁴) and local linkage disequilibrium, whereas blue points represent the remaining tested SNPs. Based on these results, subsequent interpretation was structured according to both biological trait groups and statistical evidence level, distinguishing corrected SNP associations from suggestive SNP-level associations.
Figure 2.
Manhattan plots of genome-wide SNP marker–trait associations for drought-related traits under rainfed field conditions. Manhattan plots are shown for relative water content (RWC; (A)), plant height (PH; (B)), dry weight (DW; (C)), fresh weight (FW; (D)), flag leaf length (FLL; (E)), and flag leaf width (FLW; (F)). Each point represents a SNP plotted according to its chromosomal position and −log10(P) value. Red points indicate selected peak markers, whereas blue points represent the remaining tested SNPs. Brown dashed lines indicate the suggestive threshold (p < 1 × 10⁻⁴), whereas yellow dashed lines in panels A and B indicate the Bonferroni threshold (p ≤ 2.53 × 10⁻⁶).
3.3. Marker–Trait Associations for RWC
RWC was considered a key physiological indicator of plant water status under rainfed conditions. GWAS identified one corrected significant association for RWC (Table 2). The strongest RWC-associated SNP was AX-86184518 on chromosome 2D, located at 560,005,079 bp (Figure 2A). This marker showed p = 1.21 × 10⁻⁶ and explained 11.94% of the genotypic variation. The two allele classes differed substantially in mean RWC values, with 67.75% for the major allele class and 82.00% for the minor allele class, suggesting a marked marker-associated difference in plant water status. Cross-model sensitivity analysis showed that AX-86184518 remained the top-ranked RWC association under the alternative MLM, whereas the strength of support for PH and the associations detected for the remaining traits varied between models (Supplementary Table S8). Additional suggestive RWC-associated SNPs were detected on chromosomes 1D, 5B and 3D (Table 2). These SNPs did not pass the corrected significance threshold and were therefore interpreted cautiously. Among all RWC associations detected in the present study, the chromosome 2D region tagged by AX-86184518 showed the strongest statistical evidence. Regional LD analysis showed that AX-86184518 had only weak LD with surrounding array markers within the ±5 Mb interval (maximum r2 with another SNP = 0.042; Supplementary Figure S3A), indicating that a clear local LD block could not be resolved around this marker with the present SNP dataset.
Table 2.
Selected SNP marker–trait associations detected for drought-related traits.
3.4. Marker–Trait Associations for Plant Growth and Biomass-Related Traits
Among the analyzed growth-related traits, PH showed the strongest evidence for additive marker–trait associations. At the suggestive threshold of p < 1 × 10⁻⁴, 114 SNPs were detected for PH (Supplementary Table S3). Multiple-testing correction further supported PH as the trait with the strongest GWAS evidence, with 616 SNPs passing the FDR threshold and eight SNPs passing the Bonferroni threshold. The leading PH association was detected on chromosome 2A at marker AX-158540611, located at 694,680,772 bp (Figure 2B). This marker showed p = 2.34 × 10⁻¹¹ and explained 19.40% of the genotypic variation (Table 2). The mean PH values differed strongly between allele classes, with 96.04 cm for the major allele class and 121.91 cm for the minor allele class. A second major PH-associated SNP was detected on chromosome 4A at marker AX-94514459, located at 12,472,418 bp (Figure 2B). This marker showed p = 2.81 × 10⁻¹⁰ and explained 19.33% of the genotypic variation. Additional Bonferroni-significant PH-associated SNPs were located on chromosomes 2A, 4D, 1D, 1B, and 1A, as well as in one unassigned genomic region (Table 2). Regional analysis provided clearer local LD support for the major PH loci. AX-158540611 was in LD with the nearby Bonferroni-significant marker AX-158561628 (r2 = 0.629), located approximately 204 kb away on chromosome 2A. For the chromosome 4A locus, AX-94514459 showed LD with several nearby markers, with a maximum r2 of 0.524 (Supplementary Figure S3B,C). Together, these results indicate that PH in this wheat panel was associated with multiple genomic regions with relatively strong additive effects.
In contrast, DW and FW showed high broad-sense heritability but did not reveal associations passing the applied multiple-testing thresholds. DW had four SNPs with p < 1 × 10⁻⁴. The strongest DW-associated SNP was AX-158539426 on chromosome 7A, located at 645,628,854 bp (Figure 2C). This marker showed p = 6.20 × 10⁻⁵ and explained 8.33% of the genotypic variation. Additional suggestive DW-associated SNPs were detected on chromosomes 1D and 6B (Table 2). FW showed two suggestive SNP-level associations, including AX-158539426 on chromosome 7A (Figure 2D) and Kukri_c37738_417 on chromosome 1D. AX-158539426 showed p = 5.39 × 10⁻⁵ and explained 8.46% of the genotypic variation, while Kukri_c37738_417 showed p = 7.19 × 10⁻⁵ and explained 2.26% of the genotypic variation (Table 2). Because these associations did not pass the corrected significance thresholds, they were interpreted as preliminary statistically identified marker–trait effects.
3.5. Marker–Trait Associations Suggested Additive Associations for Flag Leaf Architecture
Flag leaf architecture was evaluated using FLL and FLW. FLL showed moderate-to-high heritability and one suggestive SNP-level association with p < 1 × 10⁻⁴ (Table 2). This association corresponded to marker Kukri_c37738_417 on chromosome 1D, with p = 5.47 × 10⁻⁵ and 2.57% of the genotypic variation explained (Figure 2E). Because this marker did not pass the corrected significance threshold, the additive GWAS evidence for FLL was limited to a suggestive marker–trait association.
FLW showed the lowest heritability among the analyzed traits, but nine SNPs reached p < 1 × 10⁻⁴ (Supplementary Table S3). The strongest FLW-associated SNP was AX-108815692 on chromosome 1B, located at 338,504,422 bp, with p = 1.14 × 10⁻⁵ and 9.74% of the genotypic variation explained (Figure 2F). Additional suggestive FLW-associated SNPs were detected on chromosomes 1B, 1D, and 4A. Among these, AX-95120043 on chromosome 1B explained 9.58% of the genotypic variation, wsnp_Ex_c1429_2745237 on chromosome 1D explained 9.16%, Excalibur_s113941_196 on chromosome 1D explained 9.07%, and Kukri_c31554_437 on chromosome 4A explained 9.00% (Table 2). Although these markers did not pass the corrected significance threshold, they represent preliminary statistically identified marker–trait associations with relatively high explained genotypic variation. Overall, flag leaf architecture traits showed less corrected additive GWAS evidence than PH and RWC, suggesting a more complex or environmentally sensitive genetic architecture.
3.6. Putative Epistatic Marker Interactions Across Drought-Related Traits
Putative epistatic interaction analysis identified marker-by-marker interactions for all six drought-related traits. After collapsing reciprocal duplicate pairs, the number of unique epistatic marker pairs differed among traits, with 755 pairs for RWC, 754 for DW, 752 for FW, 317 for FLL, 662 for FLW, and 1454 for PH (Supplementary Table S4). Following trait-specific Bonferroni correction based on the total number of unique marker-pair comparisons after pre-screening, 63 RWC, 625 DW, 115 FW, 6 FLL, 123 FLW, and 1454 PH interactions in Supplementary Table S4 passed the corresponding corrected significance threshold. For phenotypic interpretation, allele-combination means were calculated for the representative marker pairs retained in Table 3. Five representative Bonferroni-supported interactions per trait are presented in Table 3, while detailed allele-combination means and genotype-class sizes for these representative interactions are provided in Supplementary Table S5. Because some two-locus genotype classes were represented by relatively few genotypes, particularly for PH and FLW, the corresponding allele-combination means were interpreted descriptively and with caution.
Table 3.
Representative Bonferroni-supported putative epistatic marker interactions and allele-combination means for the evaluated traits.
For RWC, one representative interaction was detected between TGWA25K-TG0299 on chromosome 3B and TA009869-0749 on chromosome 5D. Across the five representative RWC interactions, allele-combination means ranged from 59.25% to 76.53%, while the within-pair contrasts ranged from 9.07 to 13.43 percentage points. PH showed the largest number of Bonferroni-supported interactions among the traits represented in Supplementary Table S4. One representative PH interaction involved Tdurum_contig42008_7461 on chromosome 5D and AX-109475947 in an unassigned genomic region. Across the five representative PH interactions, allele-combination means extended from 88.58 to 130.17 cm, with within-pair contrasts of 34.09–40.40 cm (Table 3). These phenotypic contrasts should be interpreted cautiously because some two-locus genotype classes contained relatively few genotypes.
Biomass-related traits also showed putative interaction patterns. For DW, one representative interaction was detected between RAC875_c32452_55 on chromosome 1A and RAC875_c16839_188 on chromosome 7B. Across the five representative DW interactions, allele-combination means ranged from 0.10 to 0.19 g, with within-pair contrasts of 0.07–0.08 g. For FW, one representative interaction involved RAC875_c32452_55 on chromosome 1A and AX-158578018 on chromosome 3B. In this trait, allele-combination means ranged from 0.33 to 0.71 g, with within-pair contrasts of 0.28–0.29 g (Table 3).
Flag leaf traits showed more moderate interaction-associated contrasts. One representative FLL interaction was detected between AX-94393315 on chromosome 3A and wsnp_Ex_c1943_3663067 on chromosome 5B. Across the five representative FLL interactions, allele-combination means ranged from 18.59 to 25.80 cm, with within-pair contrasts of 3.61–6.54 cm. For FLW, one representative interaction involved Tdurum_contig5522_455 on chromosome 5B and BS00049818_51 on chromosome 6D; allele-combination means ranged from 1.60 to 1.86 cm, with contrasts of 0.25–0.26 cm (Table 3). Because some FLW genotype-combination classes were sparsely represented, these phenotypic contrasts should be regarded as descriptive. Overall, the Bonferroni reassessment identified statistically supported marker-by-marker interactions across all six traits; however, these interactions are considered putative because statistical interaction does not by itself establish biological epistasis and the analysis was conducted in a single field environment.
3.7. Candidate Gene Localization Across GWAS-Associated Genomic Regions
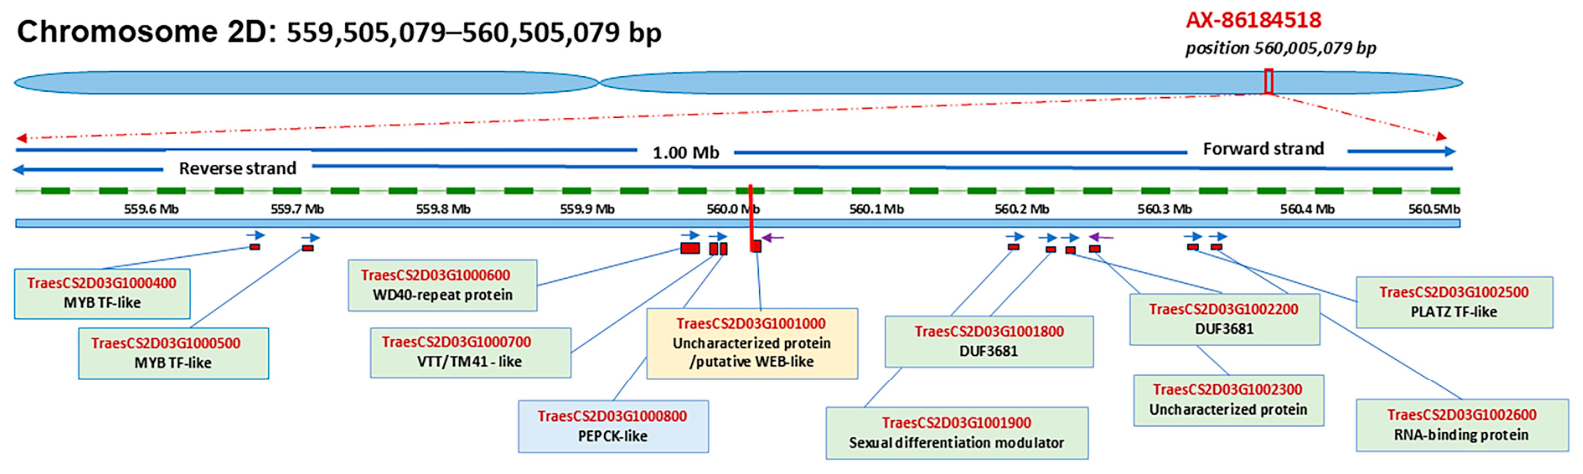
Candidate gene localization was performed for GWAS-associated genomic regions identified for the evaluated drought-related traits. Within the ±500 kb physical interval centered on the Bonferroni-significant RWC-associated SNP AX-86184518 (2D: 560,005,079 bp), 12 non-redundant high-confidence genes were identified (Figure 3; Supplementary Table S6). AX-86184518 was located in an intergenic region between TraesCS2D03G1000800 and TraesCS2D03G1001000. The closest gene was TraesCS2D03G1001000, with the nearest gene boundary located only 291 bp from the SNP position. This gene was predicted to encode a 795-aa protein corresponding to UniProt A0A1D5UM13 and RefSeq protein XP_044334216.1. Although annotated as an uncharacterized protein, family-domain evidence indicated similarity to a chloroplastic WEB family-like protein. NCBI BLASTp analysis (https://blast.ncbi.nlm.nih.gov/Blast.cgi, accessed on 11 September 2026) [45] revealed highly conserved cereal homologues, including a Triticum turgidum subsp. durum protein with 97% identity, 98% positives, and an E-value of 0.0. In silico expression profiling of the RefSeq v1.1 counterpart TraesCS2D02G446900 showed detectable expression in both non-stress and abiotic-stress-associated samples, with higher aggregated mean TPM under abiotic conditions than under non-abiotic conditions (12.75 vs. 8.01 TPM; Supplementary Table S7). The neighbouring TraesCS2D03G1000800 gene, located approximately 3.5 kb from AX-86184518, encoded a PEPCK-like protein. Based on physical proximity, conserved WEB family-like features, and detectable expression under abiotic-stress-related conditions, TraesCS2D03G1001000 was retained as the most proximal candidate gene for the RWC-associated 2D locus. For PH, candidate gene localization was performed for physically anchored Bonferroni-significant loci. The two closely positioned markers on chromosome 2A, AX-158540611 and AX-158561628, were treated as a single major QTL region containing four non-redundant high-confidence genes. AX-158540611 was located approximately 4.6 kb from TraesCS2A03G1044800, encoding a non-specific lipid-transfer protein/DIR1-like protein, whereas AX-158561628 was positioned close to TraesCS2A03G1044200, annotated as a CobW/HypB/UreG-like nucleotide-binding protein with zinc-regulated GTPase metalloprotein activator domains. The PH-associated region tagged by AX-94514459 on chromosome 4A contained 13 non-redundant genes, including amino acid transporter-like proteins, F-box proteins, lipid-transfer proteins, ribulose-phosphate 3-epimerase, cytochrome P450, pentatricopeptide repeat-containing proteins, and RNA-binding proteins. Additional PH-associated regions on chromosomes 4D, 1D, 1B, and 1A included genes related to transcriptional and post-transcriptional regulation, protein turnover, mitochondrial function, membrane transport, lipid metabolism, and stress-associated protein domains (Supplementary Table S6). The PH-associated SNP on an unassigned contig was excluded because no reliable chromosomal interval could be defined. Candidate genes for selected DW-, FW-, FLL-, and FLW-associated regions were also retrieved and are summarized in Supplementary Table S6.
Figure 3.
Candidate gene region surrounding the RWC-associated SNP AX-86184518 on chromosome 2D. The schematic shows the ±500 kb interval around the Bonferroni-significant RWC-associated SNP AX-86184518 (2D: 559,505,079–560,505,079 bp) based on the IWGSC RefSeq v2.1 wheat genome annotation. The red vertical line marks the lead SNP position at 560,005,079 bp. Gene models are displayed according to physical position and transcriptional orientation on the forward and reverse strands. Arrows indicate transcriptional orientation, and connecting lines link the gene labels to their corresponding genomic positions. TraesCS2D03G1001000, located on the reverse strand, was the closest gene to the marker, whereas TraesCS2D03G1000800 represented the nearest functionally annotated flanking gene.
4. Discussion
The present study showed that drought-related traits in the evaluated bread wheat panel were not controlled as a single uniform response but rather as partly independent physiological and morphological components. This was most clearly evident from the correlation analysis. Biomass traits were strongly associated with flag leaf morphology, whereas RWC showed only a limited association with biomass and almost no relationship with PH. This suggests that the ability to maintain leaf hydration under terminal rainfed stress was not simply a consequence of larger plant size or greater vegetative growth. Similar conclusions have been drawn from physiological studies in wheat, where drought adaptation was shown to depend on coordinated but distinct mechanisms, including water retention, osmotic adjustment, stomatal regulation, and maintenance of photosynthetic activity [10,18].
The partial independence of RWC from plant height and dry biomass is important for interpreting the GWAS results. RWC is commonly considered a direct indicator of leaf water status, but it is not necessarily expected to follow the same genetic architecture as growth-related traits. Condorelli et al. [20] showed in durum wheat that osmotic adjustment, RWC, and chlorophyll retention were physiologically connected under drought; however, their genetic control was distributed across several genomic regions. Similarly, Czyczyło-Mysza et al. [46] showed that traits related to excised-leaf water loss are genetically complex and may overlap with leaf morphology without being simply determined by leaf size. The strongest association for RWC was detected on chromosome 2D at AX-86184518. This locus is of particular interest because it passed the corrected significance threshold under rainfed field conditions, where physiological traits are usually affected by environmental heterogeneity. Previous studies have reported loci associated with RWC or water-status-related traits on chromosomes 5A and 5D in biparental wheat material [47] and drought-related loci on chromosome 2A in association-mapping populations [48]. The present lead marker, AX-86184518, is located on chromosome 2D at 560.0 Mb and therefore represents a genomic position distinct from the loci reported in these studies. Therefore, AX-86184518 represents the strongest RWC-associated marker identified in this panel under the evaluated field conditions. Accordingly, AX-86184518 should be regarded as a putative candidate marker whose stability requires validation in independent populations and across additional environments. The higher RWC associated with this locus should be interpreted as improved maintenance of leaf water status rather than as direct evidence of enhanced drought tolerance. Higher RWC under water-limited conditions may result from several physiological mechanisms, including improved water retention, reduced transpirational water loss, stomatal regulation, or osmotic adjustment [10,20]. However, these mechanisms were not directly measured in the present study; therefore, the physiological basis of the AX-86184518 association cannot yet be distinguished. Its contribution to drought adaptation should be further evaluated together with additional physiological and agronomic traits, including osmotic adjustment, stomatal conductance, canopy temperature, root characteristics, and grain yield. This is relevant because RWC is strongly influenced by developmental stage, stress intensity, and genetic background, as also shown in recent GWAS studies of wheat under drought, osmotic, and salt-related stress [25,49,50]. The candidate-gene content around AX-86184518 gives this locus additional biological interest. The nearest gene, TraesCS2D03G1001000, is annotated as an uncharacterized WEB-family-like protein. WEB/PMI2-related proteins have been studied in Arabidopsis as components involved in blue-light-induced chloroplast photorelocation [51,52]. This provides a plausible connection with chloroplast-associated stress responses. Under terminal drought, leaves are exposed not only to water deficit but also to high irradiance and oxidative pressure; therefore, genes related to chloroplast positioning or chloroplast protection represent biologically relevant positional candidates near a water-status locus. The neighbouring gene TraesCS2D03G1000800, annotated as a PEPCK-like protein, is also physiologically relevant. PEPCK-related enzymes participate in organic acid metabolism and carbon remobilization, processes that may influence source–sink balance and stress responses [53,54]. Taken together, the positional evidence, sequence conservation, in silico expression profile, and previous functional information on WEB/PMI2-related proteins support TraesCS2D03G1001000 as a prioritized positional candidate at the chromosome 2D RWC locus. However, these lines of evidence do not establish a causal role in RWC regulation, which will require direct functional validation. PH showed a much clearer additive genetic pattern than the other traits. Several markers passed the corrected threshold, especially on chromosomes 2A and 4A, indicating that PH was under strong genetic control in this panel. This is consistent with previous wheat studies, where PH has repeatedly been shown to be a highly heritable trait controlled by loci affecting internode elongation, semi-dwarfing response, and overall plant architecture [11,12,14]. In the present study, PH was not significantly correlated with RWC, biomass traits, or flag leaf dimensions. This suggests that, in this germplasm, height variation was largely independent from leaf hydration and vegetative biomass formation. For breeding under rainfed conditions, this is useful because plant architecture may be adjusted without necessarily changing water-status traits. The very high H2 estimated for PH (0.991) should be interpreted in the context of the single-environment design. Because G × E variance could not be estimated, this value may be higher than an across-environment heritability estimate, and multi-environment evaluation is required to assess its stability. The detected PH loci on 2A and 4A are also interesting in relation to previous mapping studies. Chromosome 2A has repeatedly been implicated in plant-height variation in wheat, including recent analyses of PH-related traits [11]. Chromosome 4A has also appeared in studies of drought- and heat-adaptive traits, where QTLs for canopy temperature and yield-related responses under stress were reported [55]. The major PH loci identified in the present study did not overlap known major dwarfing genes; the 4A locus at 12.47 Mb is distant from Rht-A1, while the 2A locus at 694.68 Mb is distinct from the previously reported Rht7 region. Therefore, the present results support the general importance of these chromosomes for wheat architecture and stress-adaptive plant development.
Biomass traits showed a different pattern. Although fresh and dry weight were closely correlated and had relatively high heritability, they did not produce additive associations passing the corrected multiple-testing thresholds. This suggests that biomass accumulation under terminal drought in this panel may not be controlled by one or two large-effect loci. Instead, it may depend on many small-effect loci, environment-sensitive effects, or non-additive interactions. This agrees with the broader GWAS literature on wheat drought response, where terminal drought traits at the reproductive stage often show complex and polygenic inheritance [25,50]. The shared suggestive association on chromosome 7A for FW and DW is therefore worth noting as a putative genomic region contributing to biomass accumulation under rainfed stress.
Flag leaf morphology was strongly associated with biomass at the phenotypic level. This is expected, because the flag leaf is an important photosynthetic source during reproductive development and grain filling. Yang et al. [15] showed that flag leaf length, width, and area are quantitatively inherited and influenced by water regime, whereas Wang et al. [16] reported that flag-leaf-related loci may affect yield-related traits. However, in the present study, the GWAS evidence for flag leaf traits remained mainly suggestive, supporting the view that flag leaf architecture is controlled by multiple loci and is sensitive to environmental conditions.
The marker Kukri_c37738_417 on chromosome 1D (3.17 Mb) was detected as a shared suggestive association for RWC, FW, DW, and FLL. The same marker has previously been reported in a wheat GWAS for glume color rather than for drought-related traits [56]. Thus, this previous association does not provide direct functional support for its involvement in the traits evaluated here. Because Kukri_c37738_417 did not pass either FDR or Bonferroni correction in the present study, its repeated detection across traits should be regarded as a preliminary observation requiring independent validation.
The epistatic analysis provided exploratory evidence for putative non-additive marker effects across the evaluated traits. A subset of the retained marker-by-marker interactions remained statistically supported after stringent trait-specific Bonferroni reassessment. Nevertheless, these results should be interpreted cautiously. Statistical marker-by-marker interactions do not necessarily represent direct biological interactions, and some allele-combination classes were represented by relatively few genotypes. Moreover, the analysis was conducted in a single field environment. Therefore, the observed interactions should be regarded as putative rather than as an explanation for the absence of strong additive associations. Validation in independent populations and across multiple environments will be required to establish their reproducibility and biological relevance.
The present GWAS was conducted using a relatively limited panel of 187 genotypes evaluated at a single location during one growing season. Although this population size was sufficient to detect several moderate- to large-effect associations, the statistical power to detect minor-effect loci underlying complex drought-related traits may have been limited. Moreover, the detected marker–trait associations should be interpreted within the environmental conditions of this field trial. Their stability across years and environments remains to be established. Multi-environment evaluation and validation in independent populations will therefore be required to distinguish environmentally stable genetic effects from environment-specific associations and genotype-by-environment interactions.
5. Conclusions
Overall, the study reveals three main genetic patterns. First, RWC was associated with a putative candidate locus on chromosome 2D, with AX-86184518 representing the strongest RWC-associated marker identified in this panel under the evaluated field conditions. Second, PH was controlled by several strong additive loci and was largely independent from RWC and biomass traits. Third, biomass and flag leaf traits were phenotypically connected but genetically more complex, with mainly suggestive additive markers and evidence of putative epistatic effects. These results are consistent with the current understanding of drought adaptation in wheat as a multi-component trait, where physiological water status, canopy architecture, and biomass formation should be evaluated together rather than reduced to a single marker or trait. Future studies should validate the stability of the 2D RWC locus and the major PH-associated loci in independent populations and across multiple years, locations, and drought scenarios, and should functionally characterize the prioritized candidate genes. Integrating validated loci with grain yield, canopy temperature, osmotic adjustment, root traits, and high-throughput phenotyping indices will help to clarify their contribution to drought adaptation and assess their potential breeding value.
Supplementary Materials
The following supporting information can be downloaded at: https://www.mdpi.com/article/10.3390/applbiosci5030083/s1, Figure S1: Representative field view of the bread wheat diversity panel grown under rainfed conditions at the Gobustan Regional Experimental Station during the 2022/2023 growing season; Figure S2: Quantile–quantile (QQ) plots of genome-wide association analysis for relative water content (RWC), plant height (PH), dry weight (DW), fresh weight (FW), flag leaf length (FLL), and flag leaf width (FLW). Blue points represent the observed −log10(P) values plotted against the expected −log10(P) values under the null hypothesis, and the diagonal line represents the expected distribution; Figure S3: Regional association plots and pairwise linkage disequilibrium (LD) heatmaps for the major RWC-associated locus AX-86184518 on chromosome 2D and the major PH-associated loci AX-158540611 on chromosome 2A and AX-94514459 on chromosome 4A. In the regional association plots, blue points represent SNPs and orange stars indicate the lead SNPs; the horizontal lines indicate the suggestive (p < 1 × 10⁻⁴) and Bonferroni (p ≤ 2.53 × 10⁻⁶) thresholds. Pairwise LD is expressed as r2, with the color scale ranging from low to high LD; Table S1: List of bread wheat genotypes used in this study, including country of origin and varietal type; Table S2: Hydro-meteorological data of the Gobustan station for the 2022/2023 growing season; Table S3: SNP marker–trait associations reaching the suggestive threshold of p < 1 × 10⁻⁴ for the six evaluated traits; Table S4: Reciprocal-collapsed putative epistatic marker interactions with trait-specific Bonferroni reassessment; Table S5: Allele-combination means and class sizes for the representative putative epistatic interactions shown in Table 3; Table S6: Putative candidate genes located within the predefined positional intervals surrounding selected GWAS lead SNPs/QTL regions, based on IWGSC RefSeq v2.1 annotation; Table S7: In silico expression summary of TraesCS2D03G1001000 and its homoeologs using Wheat Expression Browser data, including aggregated abiotic-stress-associated and non-abiotic expression profiles and condition-level abiotic expression summaries; Table S8: Cross-model sensitivity analysis of selected marker–trait associations using GRAMMAR and an alternative kinship-based mixed linear model (MLM); Table S9: Phenotypic measurements for relative water content, plant height, fresh weight, dry weight, flag leaf length, and flag leaf width across the field replications used for calculation of genotype means.
Author Contributions
I.H. supervised the project and acquired funding. S.R. and A.A.N. conceptualized the research idea. S.R., A.J., U.G. and F.K. conducted the field trials. J.L., A.A.N. and S.R. performed the statistical analysis. S.R. drafted the original manuscript. I.H., J.L., A.A.N. and S.R. contributed to the review and editing process. All authors have read and agreed to the published version of the manuscript.
Funding
This study was supported by the Azerbaijan National Academy of Sciences (2022) and by the Ministry of Science and Education of the Republic of Azerbaijan (2023).
Institutional Review Board Statement
Not applicable.
Informed Consent Statement
Not applicable.
Data Availability Statement
The data that support the findings of this study are available from Dr. Samira Rustamova (s.rustamova@imbb.science.az) upon request.
Acknowledgments
OpenAI ChatGPT (web application, https://chatgpt.com/, accessed on 11 September 2026) and Google NotebookLM (web application, https://notebooklm.google.com/, accessed on 11 September 2026) were used to support English-language editing, improve clarity and academic style, organize selected information from the literature, and refine the structure of selected sections of the manuscript. These tools were not used to generate or analyze experimental data, perform statistical analyses, produce research results, or determine the scientific conclusions. All AI-assisted content was critically reviewed, verified against the original sources, and revised by the authors, who take full responsibility for the final content of the manuscript.
Conflicts of Interest
The authors declare no conflicts of interest.
Abbreviations
The following abbreviations are used in this manuscript:
| ANOVA | Analysis of variance |
| BLASTP | Basic Local Alignment Search Tool for proteins |
| Chr | Chromosome |
| CV | Coefficient of variation |
| DW | Dry weight |
| FDR | False discovery rate |
| FLL | Flag leaf length |
| FLW | Flag leaf width |
| FW | Fresh weight |
| GWAS | Genome-wide association study |
| H2 | Broad-sense heritability |
| HC | High-confidence |
| LD | Linkage disequilibrium |
| MAF | Minor allele frequency |
| NCBI | National Center for Biotechnology Information |
| PH | Plant height |
| PVE | Percentage of genotypic variation explained |
| QTL | Quantitative trait locus |
| RWC | Relative water content |
| SNP | Single-nucleotide polymorphism |
| TPM | Transcripts per million |
| TW | Turgid weight |
References
- Poole, N.; Donovan, J.; Erenstein, O. Viewpoint: Agri-nutrition research: Revisiting the contribution of maize and wheat to human nutrition and health. Food Policy 2021, 100, 101976. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- FAO. FAOSTAT: Crops and Livestock Products; Food and Agriculture Organization of the United Nations: Rome, Italy, 2026. [Google Scholar]
- IPCC. Climate Change 2023: Synthesis Report; Intergovernmental Panel on Climate Change: Geneva, Switzerland, 2023. [Google Scholar]
- He, Y.; Zhao, Y.; Sun, S.; Fang, J.; Zhang, Y.; Sun, Q.; Liu, L.; Duan, Y.; Hu, X.; Shi, P. Global warming determines future increase in compound dry and hot days within wheat growing seasons worldwide. Clim. Change 2024, 177, 70. [Google Scholar] [CrossRef] [Scilit]
- Dadrasi, A.; Chaichi, M.; Nehbandani, A.; Soltani, E.; Nemati, A.; Salmani, F.; Heydari, M.; Yousefi, A.R. Global insight into understanding wheat yield and production through agro-ecological zoning. Sci. Rep. 2023, 13, 15898. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Farooq, M.; Hussain, M.; Siddique, K.H.M. Drought stress in wheat during flowering and grain-filling periods. Crit. Rev. Plant Sci. 2014, 33, 331–349. [Google Scholar] [CrossRef] [Scilit]
- Isgandarova, T.Y.; Rustamova, S.M.; Aliyeva, D.R.; Rzayev, F.H.; Gasimov, E.K.; Huseynova, I.M. Antioxidant and ultrastructural alterations in wheat during drought-induced leaf senescence. Agronomy 2024, 14, 2924. [Google Scholar] [CrossRef] [Scilit]
- Turner, N.C. Techniques and experimental approaches for the measurement of plant water status. Plant Soil 1981, 58, 339–366. [Google Scholar] [CrossRef] [Scilit]
- Sewore, B.M.; Abe, A.; Nigussie, M. Evaluation of bread wheat (Triticum aestivum L.) genotypes for drought tolerance using morpho-physiological traits under drought-stressed and well-watered conditions. PLoS ONE 2023, 18, e0283347. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Guizani, A.; Askri, H.; Amenta, M.L.; Defez, R.; Babay, E.; Bianco, C.; Rapanà, N.; Finetti-Sialer, M.; Gharbi, F. Drought responsiveness in six wheat genotypes: Identification of stress resistance indicators. Front. Plant Sci. 2023, 14, 1232583. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Zhang, H.; Li, Y.; Wei, N.; Hao, Y.; Li, X.; Wu, B.; Zheng, X.; Zhao, J.; Zheng, J. Genetic dissection of plant height-related traits by combined methods in wheat (Triticum aestivum L.). BMC Plant Biol. 2025, 25, 988. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Rebetzke, G.J.; Richards, R.A. Gibberellic acid-sensitive dwarfing genes reduce plant height to increase kernel number and grain yield of wheat. Aust. J. Agric. Res. 2000, 51, 235–245. [Google Scholar] [CrossRef] [Scilit]
- Hao, Q.; Wang, D.; Xu, Y.; Zhang, H.; Wang, Y.; Zhang, X.; Wang, L. Impact of “Green Revolution” gene Rht-B1b on coleoptile length of wheat. Front. Plant Sci. 2023, 14, 1147019. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Mo, Y.; Vanzetti, L.S.; Hale, I.; Spagnolo, E.J.; Guidobaldi, F.; Al-Oboudi, J.; Odle, N.; Pearce, S.; Helguera, M.; Dubcovsky, J. Identification and characterization of Rht25, a locus on chromosome arm 6AS affecting wheat plant height, heading time and spike development. Theor. Appl. Genet. 2018, 131, 2021–2035. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Yang, D.; Liu, Y.; Cheng, H.; Chang, L.; Chen, J.; Chai, S.; Li, M. Genetic dissection of flag leaf morphology in wheat (Triticum aestivum L.) under diverse water regimes. BMC Genet. 2016, 17, 94. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Wang, Y.; Qiao, L.; Yang, C.; Li, X.; Zhao, J.; Wu, B.; Zheng, X.; Li, P.; Zheng, J. Identification of genetic loci for flag-leaf-related traits in wheat (Triticum aestivum L.) and their effects on grain yield. Front. Plant Sci. 2022, 13, 990287. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Schierenbeck, M.; Ries, D.; Maurer, H.P.; Leiser, W.L.; Würschum, T.; Philipp, N. Natural allelic variation confers diversity in the regulation of flag leaf morphology in wheat. Sci. Rep. 2024, 14, 12889. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Sallam, A.; Alqudah, A.M.; Dawood, M.F.A.; Baenziger, P.S.; Börner, A. Drought stress tolerance in wheat and barley: Advances in physiology, breeding and genetics research. Int. J. Mol. Sci. 2019, 20, 3137. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Maulana, F.; Huang, W.; Anderson, J.D.; Ma, X.-F. Genome-wide association mapping of seedling drought tolerance in winter wheat. Front. Plant Sci. 2020, 11, 573786. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Condorelli, G.E.; Newcomb, M.; Groli, E.L.; Maccaferri, M.; Forestan, C.; Babaeian, E.; Tuller, M.; White, J.W.; Ward, R.; Mockler, T.; et al. Genome-wide association study uncovers the QTLome for osmotic adjustment and related drought-adaptive traits in durum wheat. Genes 2022, 13, 293. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Kamruzzaman, M.; Beyene, M.A.; Siddiqui, M.N.; Ballvora, A.; Léon, J.; Naz, A.A. Pinpointing genomic loci for drought-induced proline and hydrogen peroxide accumulation in bread wheat under field conditions. BMC Plant Biol. 2022, 22, 584. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Sehgal, D.; Rathan, N.D.; Özdemir, F.; Keser, M.; Akin, B.; Dababat, A.A.; Koc, E.; Dreisigacker, S.; Morgounov, A. Genomic wide association study and selective sweep analysis identify genes associated with improved yield under drought in Turkish winter wheat germplasm. Sci. Rep. 2024, 14, 8431. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- International Wheat Genome Sequencing Consortium. Shifting the limits in wheat research and breeding using a fully annotated reference genome. Science 2018, 361, eaar7191. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Walkowiak, S.; Gao, L.; Monat, C.; Haberer, G.; Kassa, M.T.; Brinton, J.; Ramirez-Gonzalez, R.H.; Kolodziej, M.C.; Delorean, E.; Thambugala, D.; et al. Multiple wheat genomes reveal global variation in modern breeding. Nature 2020, 588, 277–283. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Nouraei, S.; Mia, M.S.; Liu, H.; Turner, N.C.; Yan, G. Genome-wide association study of drought tolerance in wheat (Triticum aestivum L.) identifies SNP markers and candidate genes. Mol. Genet. Genom. 2024, 299, 22. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Mosalam, M.; Nemr, R.A.; Aljabri, M.; Said, A.A.; El-Soda, M. Exploring genomic loci and candidate genes associated with drought tolerance indices in spring wheat evaluated under two levels of drought. BMC Plant Biol. 2025, 25, 408. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Mourad, A.M.I.; Sallam, A.; Eltaher, S.; Börner, A.; Moursi, Y.S. Genome-wide association study and gene network analysis of drought tolerance in wheat during early growth. Front. Plant Sci. 2026, 17, 1775894. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Red Cross Red Crescent Climate Centre. Azerbaijan: Climate Country Profile; Red Cross Red Crescent Climate Centre: The Hague, The Netherlands, 2024. [Google Scholar]
- Rustamova, S.; Shrestha, A.; Naz, A.A.; Huseynova, I. Expression profiling of DREB1 and evaluation of vegetation indices in contrasting wheat genotypes exposed to drought stress. Plant Gene 2021, 25, 100266. [Google Scholar] [CrossRef] [Scilit]
- Aliyeva, D.R.; Gurbanova, U.A.; Rzayev, F.H.; Gasimov, E.K.; Huseynova, I.M. Biochemical and ultrastructural changes in wheat plants during drought stress. Biochemistry 2023, 88, 1944–1955. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Allahverdiyev, T.I. Effect of drought stress on some biochemical and physiological parameters of bread wheat genotypes. Trans. Inst. Mol. Biol. Biotechnol. 2024, 8, 60–64. [Google Scholar] [CrossRef] [Scilit] [PubMed][Green Version]
- Allahverdiyev, T.I.; Rzayev, F.H.; Gasimov, E.K.; Huseynova, I.M. Effect of drought stress on some biochemical parameters and ultrastructure of bread wheat genotypes. Cereal Res. Commun. 2025, 53, 2373–2387. [Google Scholar] [CrossRef] [Scilit]
- Zakieh, M.; Gaikpa, D.S.; Leiva Sandoval, F.; Alamrani, M.; Henriksson, T.; Singh, P.K.; Chawade, A. Characterizing winter wheat germplasm for Fusarium head blight resistance under accelerated growth conditions. Front. Plant Sci. 2021, 12, 705006. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Zakieh, M.; Alemu, A.; Henriksson, T.; Pareek, N.; Singh, P.K.; Chawade, A. Exploring GWAS and genomic prediction to improve Septoria tritici blotch resistance in wheat. Sci. Rep. 2023, 13, 15651. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Aulchenko, Y.S.; de Koning, D.J.; Haley, C. Genomewide rapid association using mixed model and regression: A fast and simple method for genomewide pedigree-based quantitative trait loci association analysis. Genetics 2007, 177, 577–585. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Benaouda, S.; Dadshani, S.; Koua, P.; Léon, J.; Naz, A.A. Identification of QTLs for wheat heading time across multiple environments. Theor. Appl. Genet. 2022, 135, 2833–2848. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Reinert, S.; Kortz, A.; Léon, J.; Naz, A.A. Genome-wide association mapping in the global diversity set reveals new QTL controlling root system and related shoot variation in barley. Front. Plant Sci. 2016, 7, 1061. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Naz, A.A.; Reinert, S.; Bostanci, C.; Seperi, B.; Dadshani, S.; Dreisigacker, S.; Léon, J. Mining the global diversity for bioenergy traits of barley straw: Genomewide association study under varying plant water status. GCB Bioenergy 2017, 9, 1356–1369. [Google Scholar] [CrossRef] [Scilit]
- Benjamini, Y.; Yekutieli, D. The control of the false discovery rate in multiple testing under dependency. Ann. Stat. 2001, 29, 1165–1188. [Google Scholar] [CrossRef] [Scilit]
- Bolser, D.M.; Staines, D.M.; Perry, E.; Kersey, P.J. Ensembl Plants: Integrating tools for visualizing, mining, and analyzing plant genomic data. Methods Mol. Biol. 2017, 1533, 1–31. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Smedley, D.; Haider, S.; Durinck, S.; Pandini, L.; Provero, P.; Allen, J.; Arnaiz, O.; Awedh, M.H.; Baldock, R.; Barbiera, G.; et al. The BioMart community portal: An innovative alternative to large, centralized data repositories. Nucleic Acids Res. 2015, 43, W589–W598. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- The UniProt Consortium. UniProt: The universal protein knowledgebase in 2023. Nucleic Acids Res. 2023, 51, D523–D531. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Borrill, P.; Ramirez-Gonzalez, R.; Uauy, C. expVIP: A customizable RNA-seq data analysis and visualization platform. Plant Physiol. 2016, 170, 2172–2186. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Ramírez-González, R.H.; Borrill, P.; Lang, D.; Harrington, S.A.; Brinton, J.; Venturini, L.; Davey, M.; Jacobs, J.; van Ex, F.; Pasha, A.; et al. The transcriptional landscape of polyploid wheat. Science 2018, 361, eaar6089. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Altschul, S.F.; Gish, W.; Miller, W.; Myers, E.W.; Lipman, D.J. Basic local alignment search tool. J. Mol. Biol. 1990, 215, 403–410. [Google Scholar] [CrossRef] [PubMed]
- Czyczyło-Mysza, I.M.; Marcińska, I.; Skrzypek, E.; Bocianowski, J.; Dziurka, K.; Rančić, D.; Radošević, R.; Pekić-Quarrie, S.; Dodig, D.; Quarrie, S.A. Genetic analysis of water loss of excised leaves associated with drought tolerance in wheat. PeerJ 2018, 6, e5063. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Naroui Rad, M.R.; Abdul Kadir, M.; Rafii, M.Y.; Jaafar, H.Z.E.; Naghavi, M.R. Bulked segregant analysis for relative water content to detect quantitative trait loci in wheat under drought stress. Genet. Mol. Res. 2012, 11, 3882–3888. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Ahmad, M.Q.; Khan, S.H.; Khan, A.S.; Kazi, A.M.; Basra, S.M.A. Identification of QTLs for drought tolerance traits on wheat chromosome 2A using association mapping. Int. J. Agric. Biol. 2014, 16, 862–870. [Google Scholar]
- Javid, S.; Bihamta, M.R.; Omidi, M.; Abbasi, A.R.; Alipour, H.; Ingvarsson, P.K. Genome-wide association study and genome prediction of seedling salt tolerance in bread wheat (Triticum aestivum L.). BMC Plant Biol. 2022, 22, 581. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Reddy, S.S.; Saini, D.K.; Singh, G.M.; Sharma, S.; Mishra, V.K.; Joshi, A.K. Genome-wide association mapping of genomic regions associated with drought stress tolerance at seedling and reproductive stages in bread wheat. Front. Plant Sci. 2023, 14, 1166439. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Luesse, D.R.; DeBlasio, S.L.; Hangarter, R.P. Plastid movement impaired 2, a new gene involved in normal blue-light-induced chloroplast movements in Arabidopsis. Plant Physiol. 2006, 141, 1328–1337. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Kodama, Y.; Suetsugu, N.; Kong, S.G.; Wada, M. Two interacting coiled-coil proteins, WEB1 and PMI2, maintain the chloroplast photorelocation movement velocity in Arabidopsis. Proc. Natl. Acad. Sci. USA 2010, 107, 19591–19596. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Lea, P.J.; Chen, Z.H.; Leegood, R.C.; Walker, R.P. Does phosphoenolpyruvate carboxykinase have a role in both amino acid and carbohydrate metabolism? Amino Acids 2001, 20, 225–241. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Yang, X.; Chao, Q.; Gao, Z.F.; Chen, Y.X.; Liu, Y.Y.; Mei, Y.C.; Ge, X.L.; Zhang, C.Y.; Xu, J.; Yin, P.F.; et al. ZmPEPCK2 enhances nutritional quality and yield potential by synchronizing carbon and nitrogen metabolism in maize kernels. Plant Commun. 2026, 7, 101734. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Pinto, R.S.; Reynolds, M.P.; Mathews, K.L.; McIntyre, C.L.; Olivares-Villegas, J.J.; Chapman, S.C. Heat and drought adaptive QTL in a wheat population designed to minimize confounding agronomic effects. Theor. Appl. Genet. 2010, 121, 1001–1021. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Barabaschi, D.; Volante, A.; Faccioli, P.; Povesi, A.; Tagliaferri, I.; Mazzucotelli, E.; Cattivelli, L. Ancient diversity of Triticum aestivum subspecies as source of novel loci for bread wheat improvement. Front. Plant Sci. 2025, 16, 1536991. [Google Scholar] [CrossRef] [Scilit] [PubMed]

Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2026 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license.