


Abstract
Introduction: Acinetobacter baumannii is a multidrug-resistant pathogen from the ESKAPE group, associated with nosocomial infections. Its resistance to multiple antibiotics poses a global threat. The QACE protein, an efflux pump, has been identified as a key resistance mechanism, making it a promising target for the development of new antibacterial agents. Objective: Our aim is to identify low-molecular-weight compounds derived from natural products with potential inhibitory activity against the QACE protein, using virtual screening and molecular docking studies. Materials and Methods: The three-dimensional structure of QACE was retrieved from AlphaFold, followed by energy minimization and assignment of Kollman-type charges. Ligand screening was performed using the BioMX database through structural similarity analysis (Tanimoto coefficient ≥ 0.85), using ciprofloxacin as the reference compound. Selected molecules were evaluated using SwissADME to predict their pharmacokinetic properties, and three candidates with favorable profiles were chosen. Molecular docking studies were then performed using AutoDock 4 to estimate binding affinities. Results: Voacangine was the compound with the highest structural similarity, strongest binding affinity to QACE (ΔG = −6.2 kcal/mol; ki = 28.51 μM), and stable molecular interactions including hydrogen bonds and π-stacking. It showed favorable tissue distribution, low potential for CYP3A4 inhibition, and minimal predicted cardiotoxicity (hERG channel blockade). Conclusion: Voacangine emerges as a promising candidate for inhibiting the QACE efflux pump in Acinetobacter baumannii. This study highlights the value of computer-aided drug design as an effective strategy in the search for new treatments against multidrug-resistant bacteria.
1. Introduction
Antimicrobial resistance has been recognized for decades as a global public health problem, not only because of the high mortality associated with multidrug-resistant bacteria, but also due to the growing economic burden it places on healthcare systems worldwide [1].
As part of the identification of the magnitude of this problem, the World Health Organization (WHO) and the Center for Disease Control (CDC) declared the “post-antibiotic era” through various articles in which they invited all those directly and indirectly involved to raise awareness about this growing problem [2].
Building on these warnings, projections published since 2014 estimate that without urgent action, antimicrobial resistance could cause up to 10 million deaths annually by 2050 in both developed and developing countries [3].
Despite efforts in the design, discovery, and development of new antibiotics, the success rate in finding new molecular entities capable of attacking the problem has been very low [4].
Such is the case that, by 2024, only two new molecular entities (sulopenem/etzadroxil and ceftobiprole/medocaril) were approved for clinical use, and only one new molecular entity (gepotidacin) has been approved so far in 2025 [5].
It is surprising to see how bacteria have developed the ability to generate different molecular mechanisms, including proteins that have been modified [6], proteins that hydrolyze or modify antibiotics [7], or those that surprisingly and to the amazement of the scientific community behave like true efflux pumps, that is, they “expel” antibiotics from the inside of the bacteria that possess them to the extracellular part and change the permeability of the bacterial membrane [8].
Based on the need to identify the microorganisms that cause infections with different characteristics in terms of severity but with the common denominator of antimicrobial resistance, our focus is grouped under the well-known concept of the ESKAPE pathogens [9]; Acinetobacter baumannii is one of the most concerning microorganisms.
Acinetobacter baumannii is a multidrug-resistant bacterium belonging to the ESKAPE group, which has been associated with increased mortality in people who have suffered an infection caused by this microorganism. Antibacterial management has been complicated by its broad spectrum of resistance, including to carbapenems and colistin [10].
Acinetobacter baumannii infections have become a priority in the fight against multi-resistant pathogens, because mortality from infections such as sepsis, pneumonia, wound infections and genitourinary tract infections has increased from 30% to 70% due to multi-resistant strains based on the most recent reports [11].
Based on the current situation, it is necessary to search for, analyze, and, if appropriate, propose new chemical entities to combat antimicrobial resistance, specifically against Acinetobacter baumannii, a multidrug-resistant bacterium whose infections are considered of urgent and prioritized importance among the main public health problems worldwide.
Therefore, this study focuses on the in silico identification of natural product-derived compounds targeting the QACE efflux pump of Acinetobacter baumannii, aiming to contribute to the development of potential therapeutic alternatives.
2. Materials and Methods
2.1. Molecular Target Selection Criteria
Among the pharmacological targets of Acinetobacter baumannii, we find almost all types are shared by Gram-negative microorganisms. Thus, this microorganism has been combated by inhibiting the synthesis of its proteins or by inhibiting the replication of its genetic material.
It is important to highlight that the intention of having new low-molecular-weight chemical compounds should be through the exploration of new pharmacological targets that could help combat this infection.
Now, we know that we can do so by inhibiting new pharmacological targets, such as its proteins, which are responsible for providing resistance to known conventional drugs.
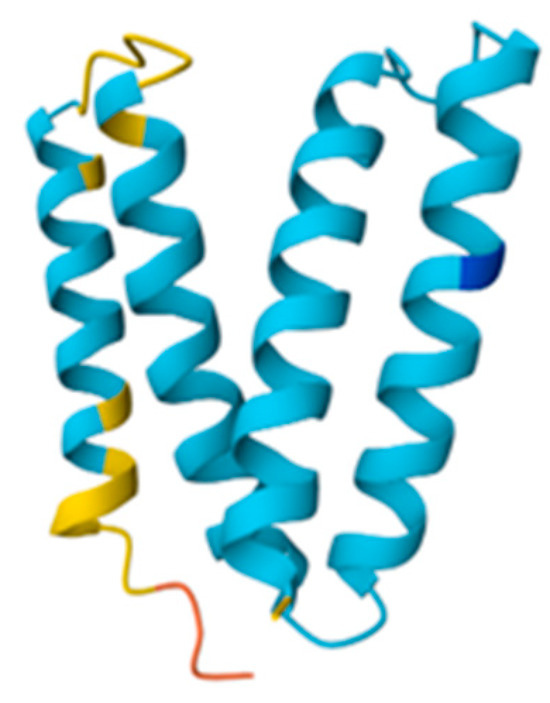
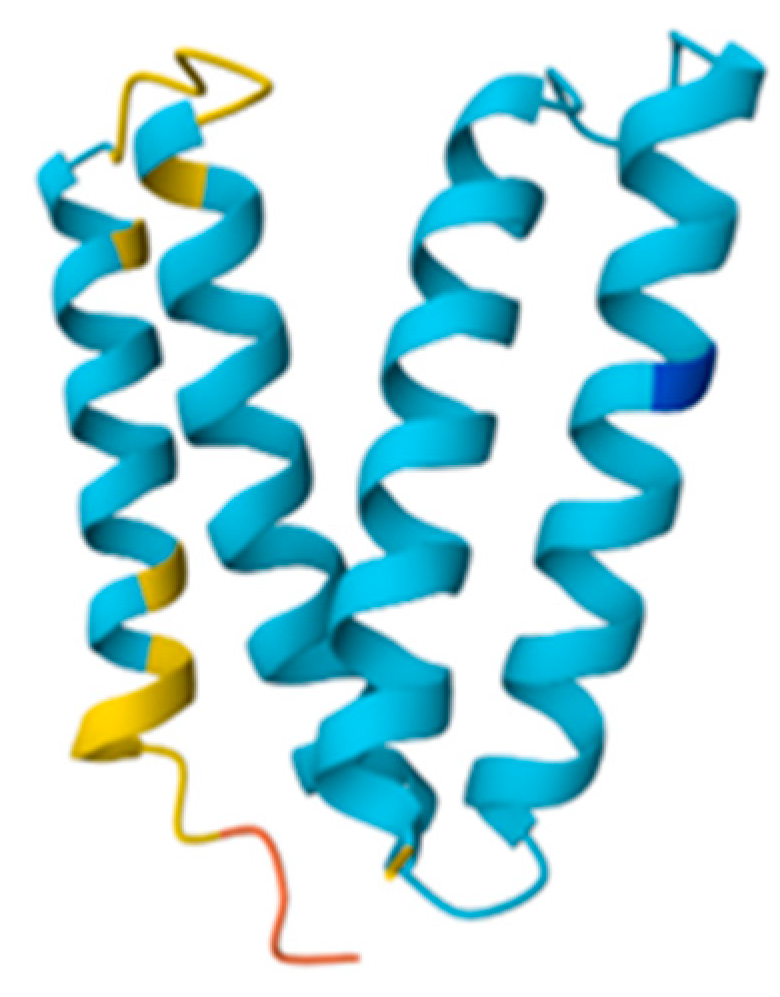
In addition to the above, Acinetobacter baumannii has several mechanisms of resistance to antibacterial agents, with efflux pumps being the most attractive for curbing its virulence; for example, the QACE protein (Figure 1), an efflux pump belonging to the QAC family of implicated in multiantibacterial resistance, was selected as a therapeutic target [12].
Figure 1.
Alphafold predicted crystallographic structure (AF-N9L2Q2-F1-v4) of the QACE receptor from Acinetobacter baumannii.
The amino acid sequence was located in the UniProt database (code N9L2Q2), and the predicted three-dimensional structure was obtained using artificial intelligence in Alphafold DB (AF-N9L2Q2-F1-v4) [12].
2.2. Virtual Screening Workflow
A search was conducted for low-molecular-weight compounds with reported antibacterial activity against similar efflux pumps (QAC type) in the ChEMBL database [13].
One hundred and ten compounds were analyzed with minimum inhibitory concentration (MIC) values using the DataWarrior software [14].
Subsequently, a structural similarity filter using the Tanimoto coefficient (0.5–0.9) was applied using the BioMX Database (from the DIFACQUIM group, UNAM) [15] to identify analogs derived from natural products.
2.3. In Silico ADMET Evaluation
Then, the selected compounds were evaluated using the SwissADME tool [16], considering parameters such as LogP, intestinal absorption, Caco-2 permeability, CYP3A4 metabolism, elimination and cardiac toxicity (hERG blockade).
The compound Voacangina was selected as the candidate with the best pharmacokinetic profile.
2.4. Molecular Docking Protocol
2.4.1. Ligand Preparation
The chemical structures obtained from BioMX Database were modeled as 2D structures using the ChemBio Ultra 12.0 software and converted into 3D structures in MDL format with their protonated states at pH = 7.4 [17].
The geometries of the compounds were calculated using the molecular mechanics force field (MMFF) in Spartan 14 [18], and, using Autodock Tools [19], the ligands were prepared by inserting polar hydrogens and Gasteiger charges in them as well as rotatable (i.e., single) bonds that were assigned by default; a PDBQT file was also generated.
2.4.2. Receptor Preparation
The X-ray coordinates of p53, p58, p38, and JNK1 were retrieved from Alphafold under the following code: AF-N9L2Q2-F1-v4; the final preparation and minimization of the receptor structures was carried out by deploying the DockPrep module of Chimera 1.15 software using the AMBER-ff14SB force field. Lastly, Kollman charges were added to the obtained structure using Autodock Tools, and a PDBQT file was generated.
2.4.3. Docking Calculations
The calculations corresponded to a rigid, blind type of molecule, carried out in Autodock4 using the Lamarckian genetic algorithm [20]. We used grid maps with 120 120 120 points in the active site of the receptor with the specific coordinates for each receptor as follows: x = −3.324, y = −0.274 and z = −0.272; a grid-point spacing of 0.375 Å was applied. AD4.dat parameters were applied to all the ligands.
The parameters used were 100 runs, a population size of 150, and a run-termination criterion of a maximum of 27,000,000 generations or a maximum of 250,000 energy evaluations. The visualization and analysis of the nonbonded interactions of the ligand–receptor interactions were visualized and analyzed using Discovery Studio Visualizer [21].
3. Results and Discussion
3.1. Molecular Target Selection
Based on the latest reports on the genomic landscape of Acinetobacter baumannii, we know that some of the genes that have a greater implication in multidrug resistance are those related to efflux pumps, where the most representative are those derived from the QAC gene (Quaternary ammonium compound-resistance protein) [22] for which there is no low-molecular-weight compound that is used clinically and/or in any clinical phase of the development of a new drug.
However, there is information on compounds for which we know their biological activity, specifically their minimum inhibitory concentration (MIC).
The first approach was to identify the QAC efflux pump gene in the Uniprot database [23]. This was followed by a selection of this gene, specifically expressed in Acinetobacter baumannii, from which the Uniprot code N9L2Q2 was obtained.
Next, a search for the crystallographic structure was carried out within the descriptive information of the receptor within the same Uniprot database. It was found that this efflux pump only has the amino acid sequence; however, the structure has not been obtained by X-ray or by new technologies such as Cryo-Electron Microscopy or Nuclear Magnetic Resonance.
3.2. Virtual Screening
At the same time, a structure-based search was performed within the ChEMBL database [13] to identify low-molecular-weight compounds that had been tested for this receptor. However, only a set of 110 low-molecular-weight compounds with experimental MIC measurements were found against a Staphylococcus aureus efflux pump expressed by the QACB gene.
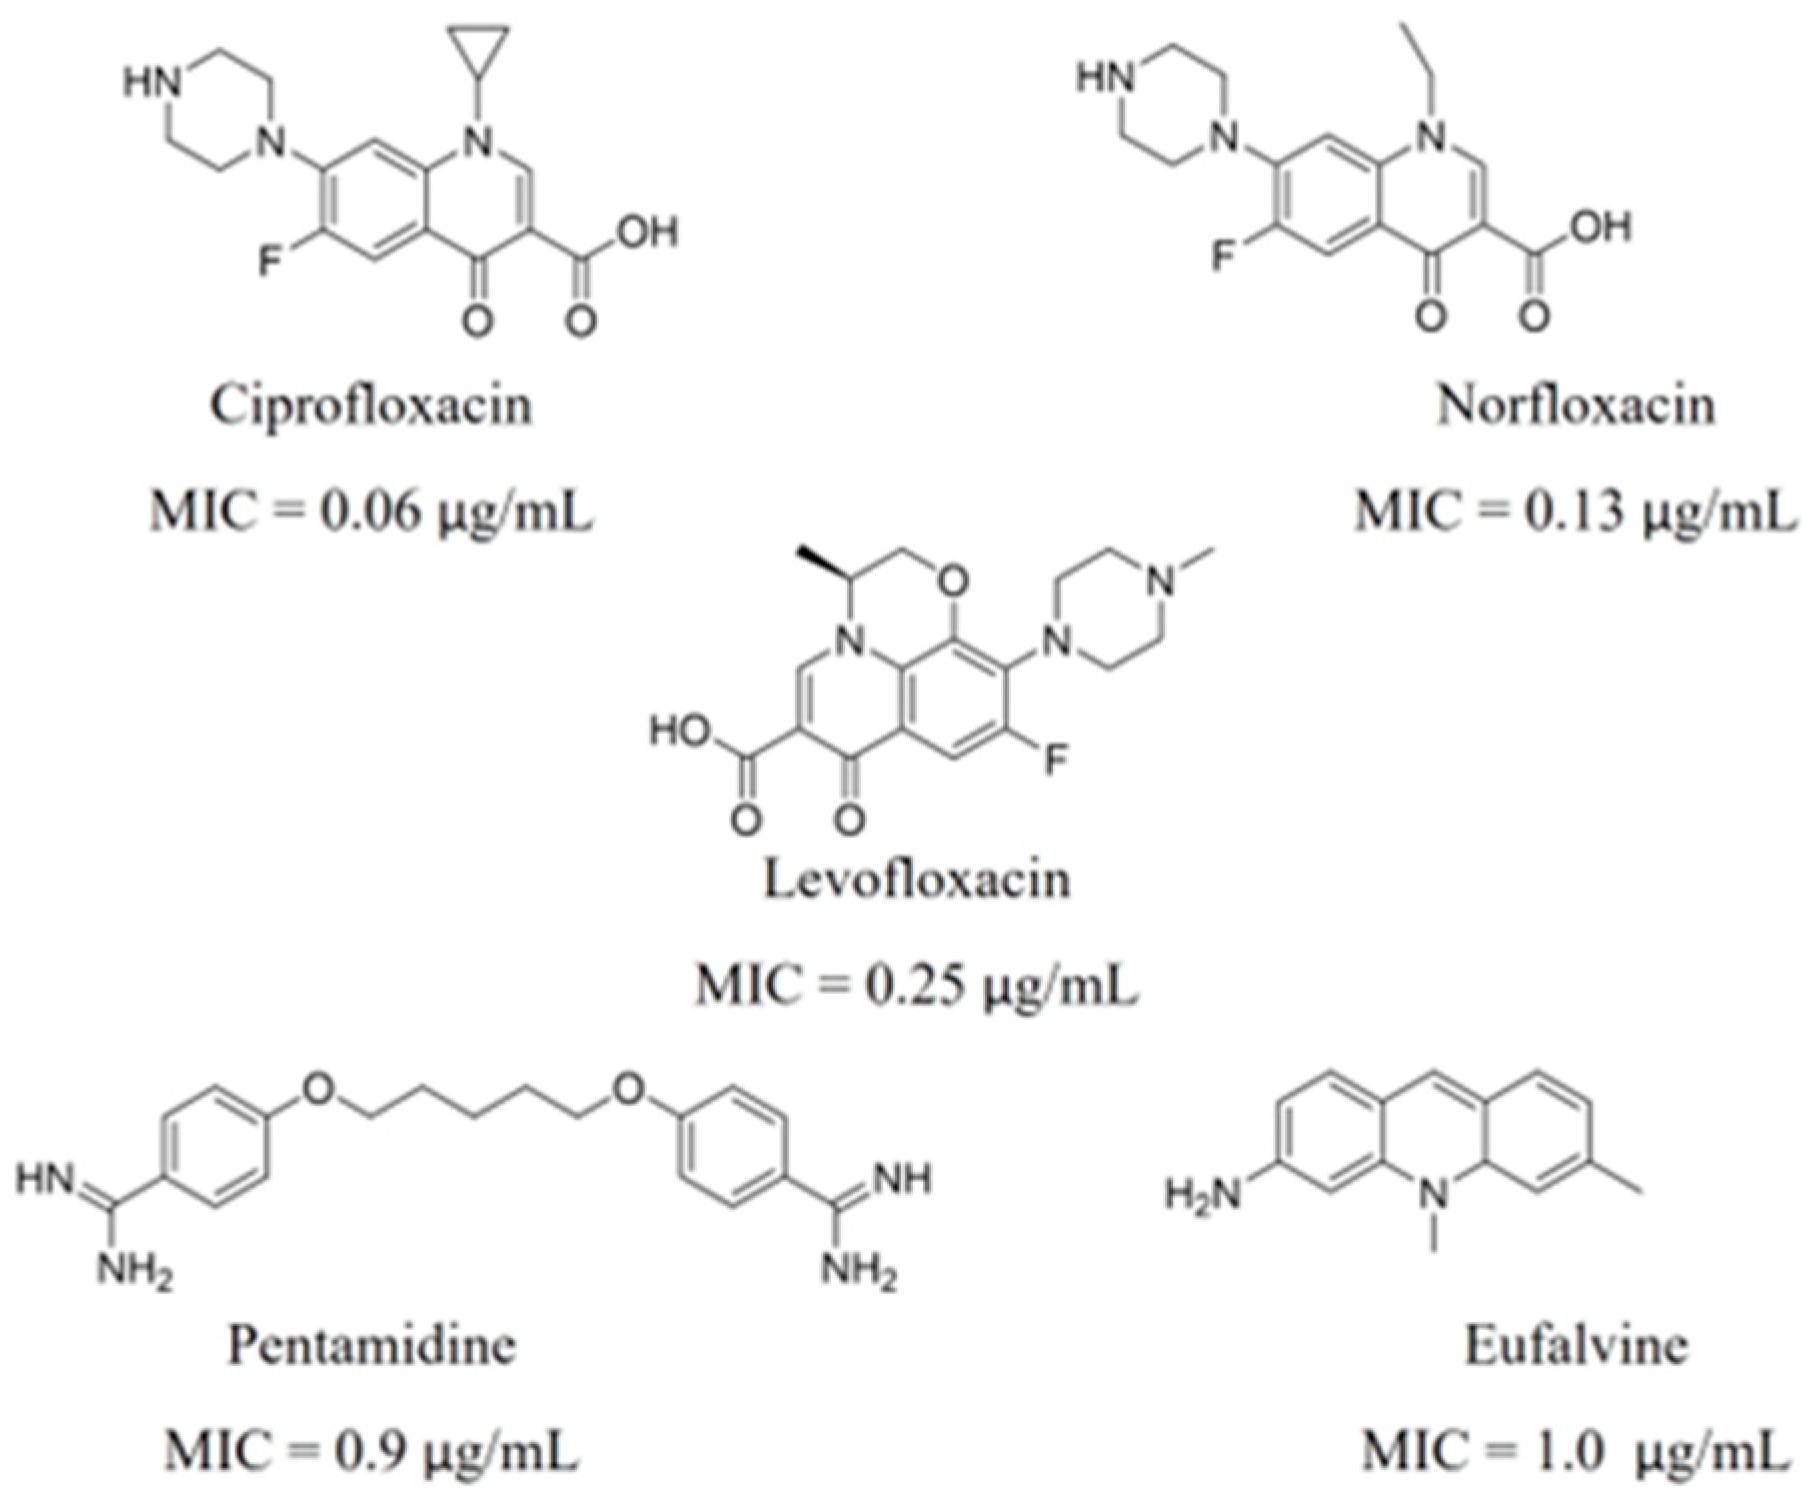
After analysis using DataWarrior [14], the top 10 low-molecular-weight compounds with the lowest MICs were selected, the drugs being presented in Figure 2.
Figure 2.
Low-molecular-weight compounds identified in ChEMBL with the best MIC against the QACB receptor.
Once the compounds with the best MIC were identified, a structural similarity search was conducted based on the Tanimoto coefficient, with a coefficient range of 0.5 to 0.9, in the BioMX database, belonging to the DIFACQUIM research group of the Faculty of Chemistry at the National Autonomous University of Mexico.
The search was carried out on a set of 605 low-molecular-weight compounds characterized and validated for their basic physicochemical characteristics.
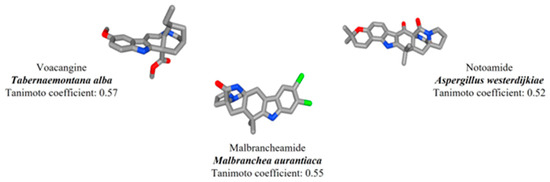
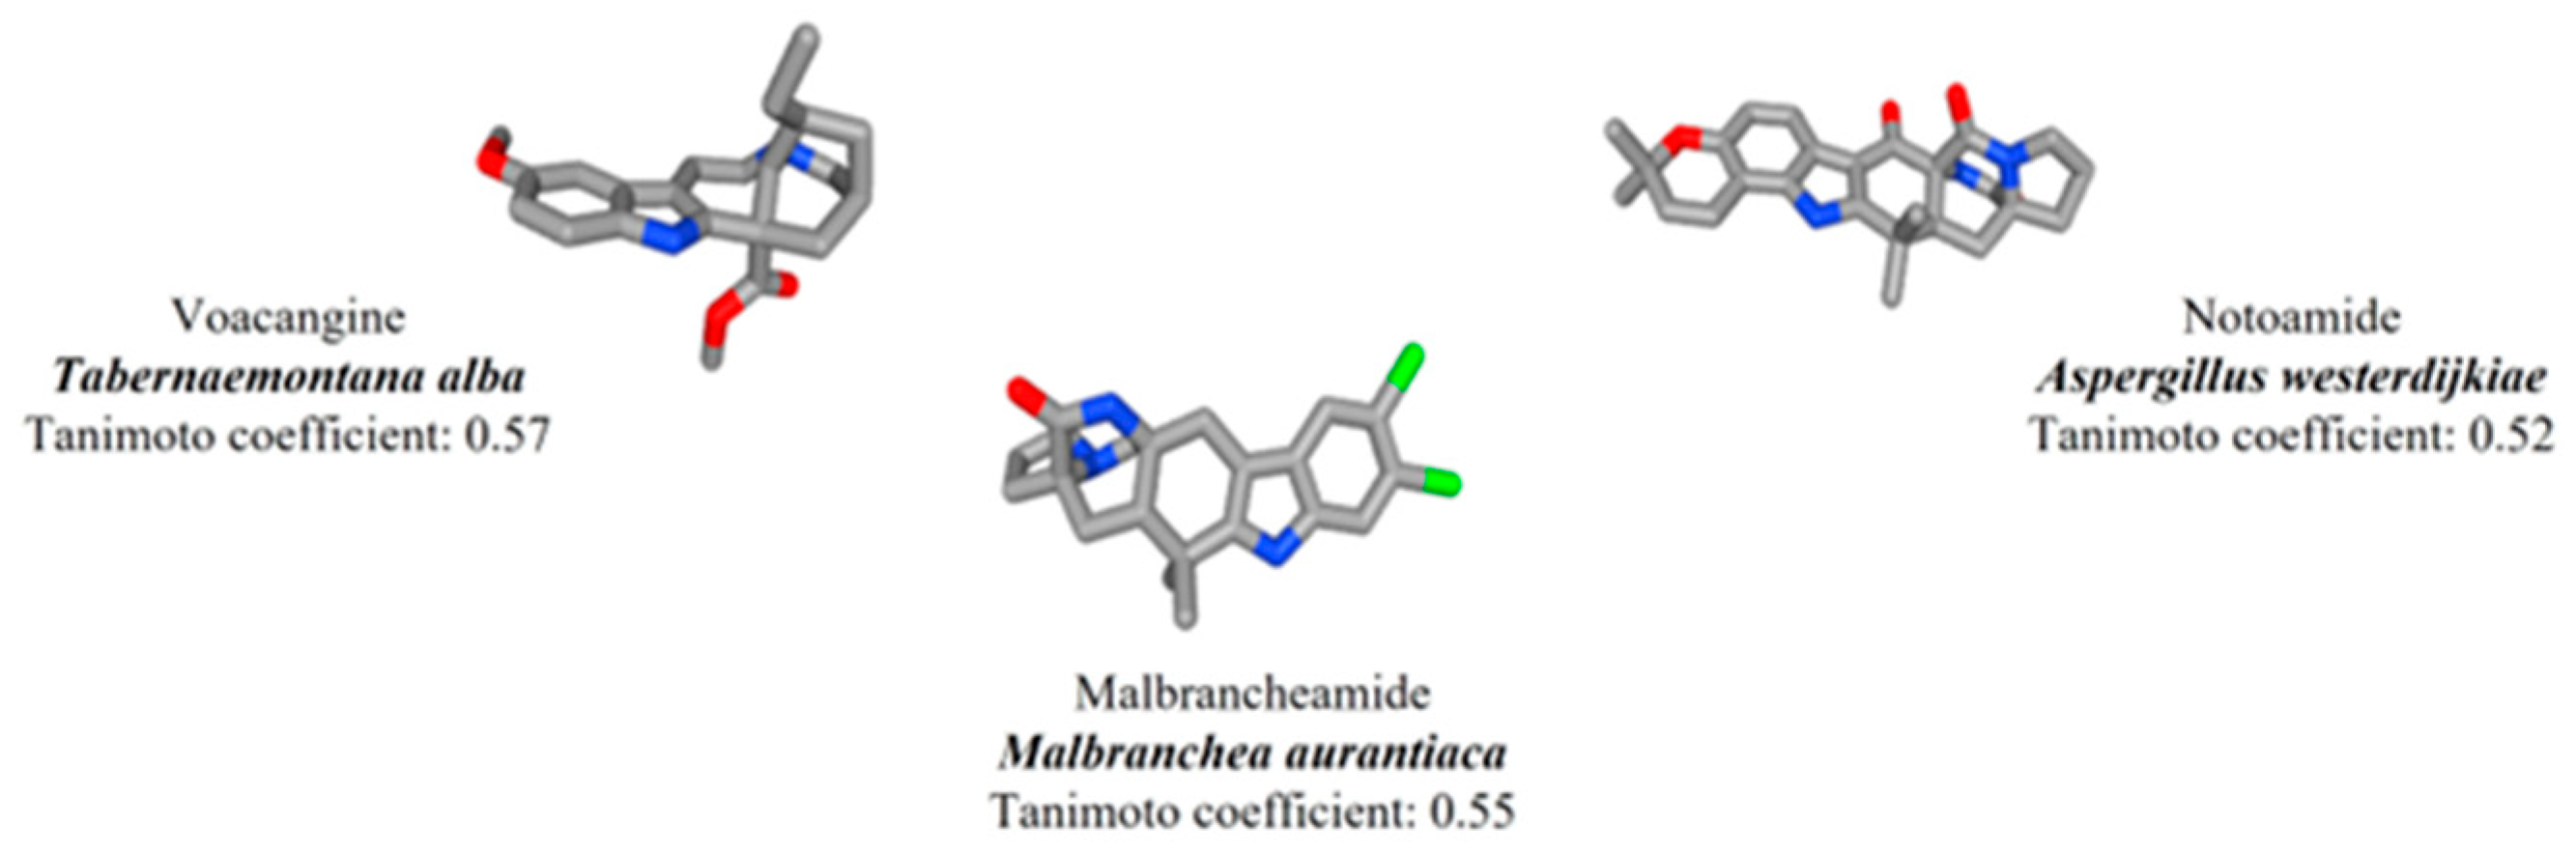
Finally, the first three low-molecular-weight compounds with the best Tanimoto coefficients were selected from the search, as shown in Figure 3.
Figure 3.
Three-dimensional schematic representation of the compounds from natural products with the best Tanimoto coefficient from the chemical similarity search based on the compounds in Figure 2.
3.3. Prediction of Pharmacokinetic Properties
Subsequently, the prediction of the pharmacokinetic properties of the three compounds was carried out in SwissADME, under the Swiss Drug Design platform and the main parameters are shown in Table 1 below.
Table 1.
Prediction of the pharmacokinetic properties of low-molecular-weight compounds found in the BioMX database.
After analyzing all the pharmacokinetic parameters, the compound with the best profile is Voacangin, due to its good absorption capacity. Furthermore, its volume of distribution is the highest among the three compounds, indicating that it can reach tissues more easily. Another advantage is that it is moderately metabolized by the CYP3A4 enzyme, which reduces the risk of rapid degradation or interaction with other medications.
It also has the highest clearance rate, which helps it be eliminated from the body without accumulating. Regarding toxicity, although it is not the least toxic, its liver and kidney values are still acceptable, and its potential to cause cardiac problems due to hERG channel blockade is relatively low.
Overall, Voacangin demonstrates a very interesting balance between efficacy and safety, making it a promising option for further investigation.
3.4. Molecular Docking Studies
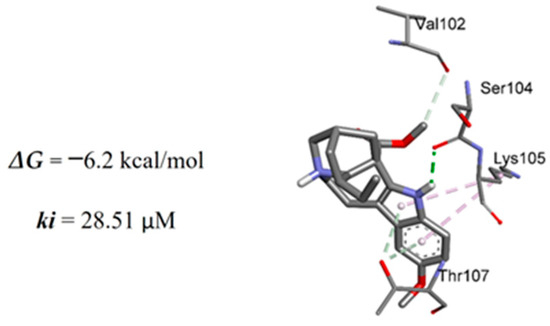
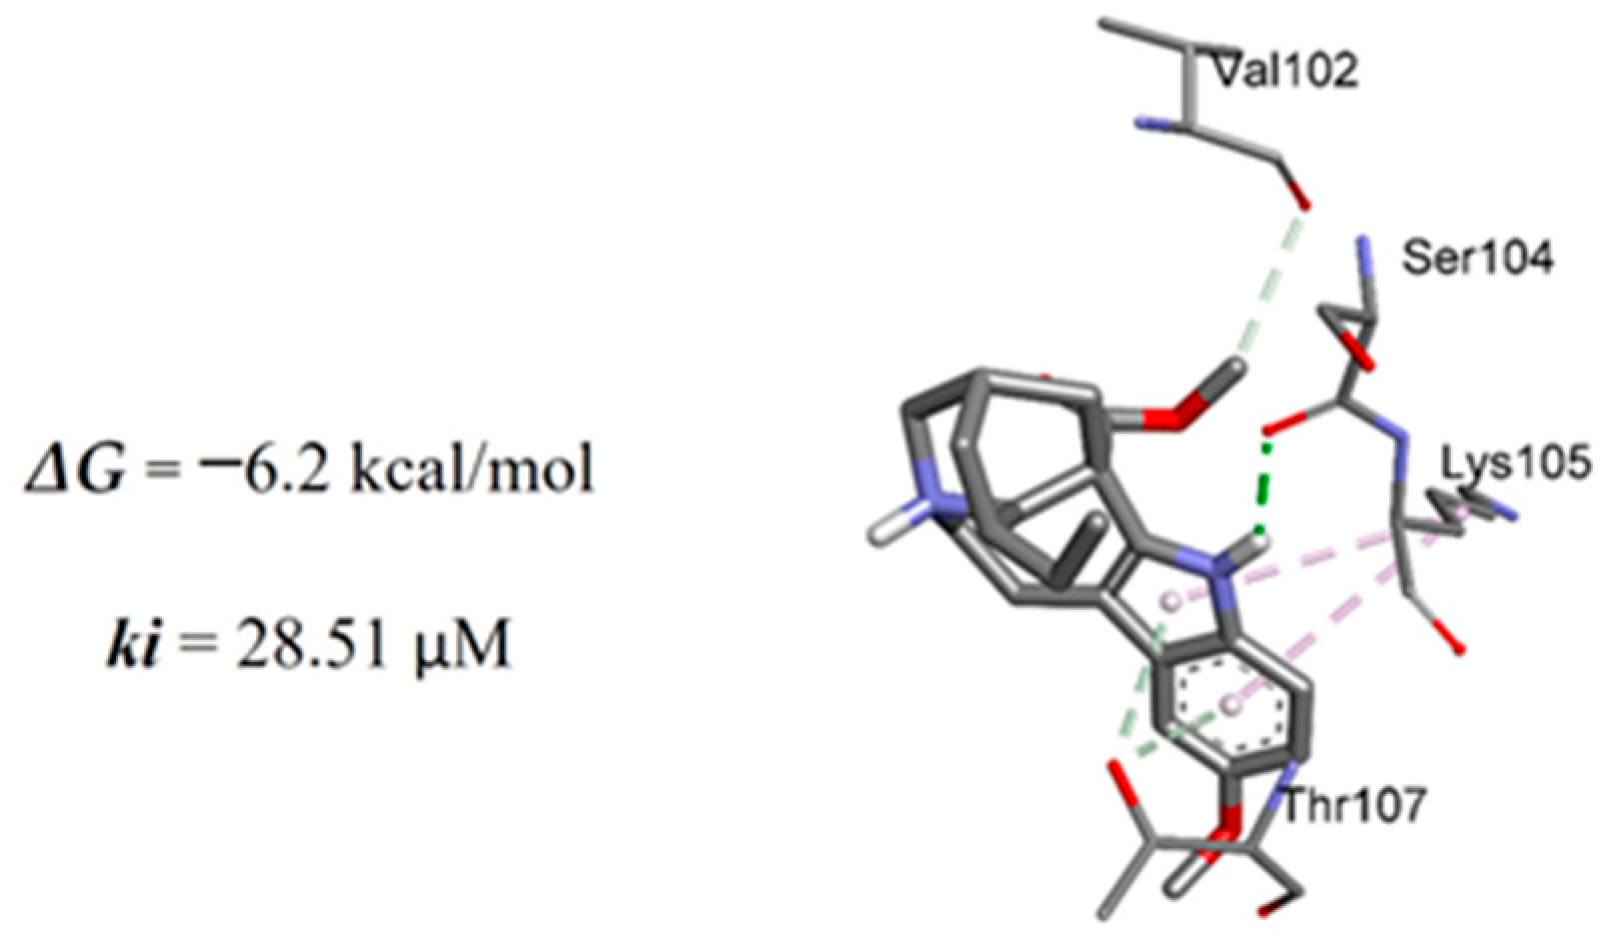
Figure 4 schematically shows the interactions between Voacangin and the QACE receptor of Acinetobacter baumannii, with an interaction free energy (ΔG) of −6.2 kcal/mol and an affinity constant (ki) of 28.51 μM.
Figure 4.
Molecular ligand–receptor interactions between Vocoangine and QACE protein of Acinetobacter baumannii.
The formation of a hydrogen bond between the available hydrogen atom of the pyrrole ring and serine 104 is noteworthy, in addition to other noncovalent interactions such as the pi-hydrophobic interactions between the indole system and Lys105.
Although we do not know the vulnerable site of the QACE receptor, we can say that it is a good start to explore the different modes of interaction of low-molecular-weight compounds with receptors of this type, which need to be explored to increase the number of possibilities of having more and better compounds with this type of biological activity.
4. Conclusions
This study identified voacangine as a natural product-derived compound with a favorable pharmacokinetic profile, low predicted toxicity, and significant binding affinity for the QACE protein of Acinetobacter baumannii.
These findings shows the usefulness of in silico approaches, such as virtual screening and molecular docking, in the strategic search for novel bioactive entities against priority pathogens, and lay the groundwork for future in vitro and in vivo validations to confirm its efficacy and safety.
Author Contributions
Conceptualization, A.S.-C.; methodology, A.S.-C. and G.F.S.-M.; software, F.R.-L.; validation, A.S.-C., G.F.S.-M. and F.R.-L.; formal analysis, A.S.-C.; investigation, G.F.S.-M. and F.R.-L.; resources, A.S.-C.; data curation, F.R.-L.; writing—original draft preparation, A.S.-C.; writing—review and editing, A.S.-C.; visualization, F.R.-L.; supervision, A.S.-C.; project administration, A.S.-C. All authors have read and agreed to the published version of the manuscript.
Funding
This research received no external funding.
Institutional Review Board Statement
Not applicable.
Informed Consent Statement
Not applicable.
Data Availability Statement
The data supporting the findings of this study are available in the ChEMBL and BioMX databases, as cited in the manuscript.
Acknowledgments
We would like to thank Julieta de la Vega Calderón, Technical Research Coordinator at the School of Medicine of the Faculty of Health Sciences at the Vasco de Quiroga University, for the facilities and support received for this work.
Conflicts of Interest
The authors declare no conflict of interest.
Abbreviations
The following abbreviations are used in this manuscript:
| ADME | Absorption, Distribution, Metabolism, Excretion |
| CDC | Centers for Disease Control and Prevention |
| ChEMBL | Chemical database of bioactive drug-like molecules |
| CYP3A4 | Cytochrome P450 3A4 |
| hERG | human Ether-à-go-go-Related Gene |
| ki | Inhibition constant |
| LogP | Octanol–water partition coefficient |
| MDL | Molecular Design Limited (chemical file format) |
| MIC | Minimum Inhibitory Concentration |
| MMFF | Merck Molecular Force Field |
| NMR | Nuclear Magnetic Resonance |
| PDBQT | Protein Data Bank, Quaternion, Torsional degrees of freedom |
| QAC | Quaternary Ammonium Compound-resistance protein |
| QACE | Quaternary Ammonium Compound-resistance Efflux protein |
| WHO | World Health Organization |
References
- Naghavi, M.; Vollset, S.E.; Ikuta, K.S.; Swetschinski, L.R.; Gray, A.P.; Wool, E.E.; Aguilar, G.R.; Mestrovic, T.; Smith, G.; Han, C.; et al. Global burden of bacterial antimicrobial resistance a systematic analysis with forecasts to 2050. Lancet 2024, 404, 1199–1226. [Google Scholar] [CrossRef] [Scilit]
- Reardon, S. WHO warns against ‘post-antibiotic’ era. Nature 2014, 15, 135–138. [Google Scholar] [CrossRef] [Scilit]
- O’Neill, J. An audience with Jim O’Neill. Nat. Rev. Drug Discov. 2016, 15, 526. [Google Scholar] [CrossRef] [Scilit]
- de la Fuente-Nunez, C.; Cesaro, A.; Hancock, R.E.W. Antibiotic failure: Beyond antimicrobial resistance. Drug Resist. Updates 2023, 71, 101012. [Google Scholar] [CrossRef] [Scilit]
- Wang, Y.; Yang, F.; Wang, B.; Xie, L.; Chen, W. New FDA drug approvals for 2024: Synthesis and clinical application. Eur. J. Med. Chem. 2025, 285, 117241. [Google Scholar] [CrossRef] [Scilit]
- Chellat, M.F.; Raguž, L.; Riedl, R. Targeting Antibiotic Resistance. Angew. Chem. Int. Ed. 2016, 55, 6600–6626. [Google Scholar] [CrossRef] [Scilit]
- Do, T.; Page, J.E.; Walker, S. Uncovering the activities, biological roles, and regulation of bacterial cell wall hydrolases and tailoring enzymes. J. Biol. Chem. 2020, 295, 3347–3361. [Google Scholar] [CrossRef] [Scilit]
- Blanco, P.; Hernando-Amado, S.; Reales-Calderon, J.A.; Corona, F.; Lira, F.; Alcalde-Rico, M.; Bernardini, A.; Sanchez, M.B.; Martinez, J.L. Bacterial Multidrug Efflux Pumps: Much More Than Antibiotic Resistance Determinants. Microorganisms 2016, 4, 14. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Miller, W.R.; Arias, C.A. ESKAPE pathogens: Antimicrobial resistance, epidemiology, clinical impact and therapeutics. Nat. Rev. Microbiol. 2024, 22, 598–616. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Arshad, N.; Azzam, W.; Zilberberg, M.D.; Shorr, A.F. Acinetobacter baumannii Complex Infections: New Treatment Options in the Antibiotic Pipeline. Microorganisms 2025, 13, 356. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Sharma, R.; Lakhanpal, D. Acinetobacter baumannii: A comprehensive review of global epidemiology, clinical implications, host interactions, mechanisms of antimicrobial resistance and mitigation strategies. Microb. Pathog. 2025, 204, 107605. [Google Scholar] [CrossRef] [Scilit]
- David, A.; Islam, S.; Tankhilevich, E.; Sternberg, M.J.E. The AlphaFold Database of Protein Structures: A Biologist’s Guide. J. Mol. Biol. 2022, 434, 167336. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Gaulton, A.; Hersey, A.; Nowotka, M.; Bento, A.P.; Chambers, J.; Mendez, D.; Mutowo, P.; Atkinson, F.; Bellis, L.J.; Cibrián-Uhalte, E.; et al. The ChEMBL database in 2017. Nucleic Acids Res. 2017, 45, D945. [Google Scholar] [CrossRef] [Scilit]
- Sander, T.; Freyss, J.; von Korff, M.; Rufener, C. DataWarrior: An Open-Source Program For Chemistry Aware Data Visualization And Analysis. J. Chem. Inf. Model. 2015, 55, 460–473. [Google Scholar] [CrossRef] [Scilit]
- Martínez-Urrutia, F.; Medina-Franco, J.L. BIOMX-DB: A web application for the BIOFACQUIM natural product database. Mol. Inform. 2024, 43, e202400060. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Daina, A.; Michielin, O.; Zoete, V. SwissADME: A free web tool to evaluate pharmacokinetics, drug-likeness and medicinal chemistry friendliness of small molecules. Sci. Rep. 2017, 7, 42717. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Milne, G.W.A. Software Review of ChemBioDraw 12.0. J. Chem. Inf. Model. 2010, 50, 2053. [Google Scholar] [CrossRef] [Scilit]
- Hehre, W.J.; Huang, W.W. Chemistry with Computation: An Introduction to SPARTAN; Wavefunction, Inc.: Irvine, CA, USA, 1995. [Google Scholar]
- Pacheco, A.B.; Hpc, L. Introduction to AutoDock and AutoDock Tools; Louisiana State University: Baton Rouge, LA, USA, 2012. [Google Scholar]
- Morris, G.M.; Goodsell, D.S.; Halliday, R.S.; Huey, R.; Hart, W.E.; Belew, R.K.; Olson, A.J. Automated docking using a Lamarckian genetic algorithm and an empirical binding free energy function. J. Comput. Chem. 1998, 19, 1639–1662. [Google Scholar] [CrossRef] [Scilit]
- Jejurikar, B.L.; Rohane, S.H. Drug designing in discovery studio. Asian J. Res. Chem. 2021, 14, 135–138. [Google Scholar]
- Pearl, S.; Anbarasu, A. Genomic landscape of nosocomial Acinetobacter baumannii: A comprehensive analysis of the resistome, virulome, and mobilome. Sci. Rep. 2025, 15, 18203. [Google Scholar] [CrossRef] [Scilit]
- The UniProt Consortium. UniProt: A hub for protein information. Nucleic Acids Res. 2015, 43, D204–D212. [Google Scholar] [CrossRef] [Scilit] [PubMed]

Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2025 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license (https://creativecommons.org/licenses/by/4.0/).