


Abstract
In this paper, we show that the sphere-in-contact model can predict long-range surface adsorption phenomena based on adsorbate-adsorbate repulsions and their geometric distance, assuming that their negative surface-induced charge is smeared on the surface of the adsorbate atoms. Additionally, it can be used to model collective surface diffusion mechanisms such as the domino-type surface diffusion of adsorbate rows on close-packed metal HCP and FCC surfaces. We have recently shown that the sphere-in-contact model can be used as an educational and research tool in various contexts, such as the visualization of carbon structures (e.g., graphene, carbon nanotubes, carbon nanocones, and graphite), heterogeneous catalysts, metal nanoparticles, and organic molecules. Here we present how it can be used to model the adsorbate structure of monoatomic elements on the hexagonal close-packed surface of HCP and FCC metals to study long-range ordering phenomena of monoatomic adsorbates on metals. We have used atoms of varying radius and color to represent the metal surface atoms and the adsorbate atoms. The study reveals that many surface configurations are possible for a fixed adsorbate coverage (θ) by the movement of the adsorbate atoms in response to surface adsorbate-adsorbate repulsions. The movement of the particles (e.g., particle diffusion) can be seen directly in the model, and this is caused by the user intervention. This has great educational and research value, as one can directly see how the adsorbate atoms reorder on the surface of a metal and therefore study diffusion mechanisms. We calculate the repulsive interaction energy of adsorbates using the sphere-in-contact model and can identify which surface-adsorbed configuration is the lowest energy. We find that at a surface coverage of 1/3 (0.333 ML), the most stable adsorbate configuration places adsorbates at the third nearest neighbor 3-fold hollow sites, forming a hexagonal pattern. We find that this model will be useful in the rational design of catalytic materials and material coatings with new technological applications where long-range ordering of surface adsorbates is essential and adsorbate interactions are mainly repulsive interatomic interactions.
1. Introduction
Rational design of catalytic surfaces is of paramount importance in heterogeneous catalysis and surface chemistry [1,2]. This is because the tailored design of adsorbate-surface systems can find applications in catalysis, nanotechnology, and materials science [3]. Particularly when these systems are composed of monoatomic adsorbates and the surface is made from transition and noble metals [4]. Therefore, a better understanding of possible surface structures of adsorbates on metal surfaces and the design of new material coatings with particular surface termination and therefore tailored properties are desirable in many fields [5]. Thus far, the understanding of adsorption on metal surfaces on the microscopic level has relied on computer models and DFT calculations, to which experimentalists lack accessibility [6]. We have previously shown that physical molecular models can have some advantages with respect to computer models, as the user can manipulate the physical molecular model in ways that are currently not possible in computer simulations [7]. Perhaps, in the future, there will be some computer models of catalytic surfaces where the structure of adsorbates can be altered during the course of the calculation by the influence of a user action. Such efforts are currently underway via virtual reality techniques [8]. Nevertheless, the use of inexpensive physical molecular models is desirable in research and education. For this reason, we have used the sphere-in-contact model to study the structure of monoatomic adsorbates on the surface of close-packed metals.
The sphere-in-contact model is a molecular model in which the atoms are represented by their atomic radius [9]. These spheres are in contact at a single point when there is a formation of a chemical bond between atoms. This model has been shown to have certain advantages over other molecular models, such as the ball-and-stick model, where the electron density volume of the molecular structure is underestimated. Also, in the wireframe model (e.g., stick model), the volume where the electron density is completely absent, we can only see the connectivity of the atoms. The other model broadly used in chemistry was developed by Robert Corey, Linus Pauling, and Walter Koltun and is called the space-filling model (also known as the CPK model) [10]. In this model, the atoms have their van der Waals (VdW) radius. This model is useful when packing and self-assembly of polycyclic aromatic hydrocarbons (PAHs) is performed, as the molecules have the correct separation due to weak interactions such as London dispersion forces [11,12]. However, when the interstitial diffusion in metals [13,14] is studied, then physical or computational molecular models using the sphere-in-contact model may provide a more accurate representation of the void space in the crystal structure, which may offer a physical picture of dopant concentration, structure, and diffusion pathways in metals for atoms such as hydrogen, carbon, and nitrogen, using geometric rules such as Hägg’s rule [15]. Such models, such as Hägg’s rule, assess the geometric space available in octahedral and tetrahedral holes of metals. Such geometric models may become an approximation for the screening of materials that then could be treated using more reliable and accurate quantum mechanical methods [16].
We have shown previously that the sphere-in-contact model can be used in research, for example, to understand the structure of carbon on curved surfaces, such as the hemispherical cap of carbon nanotubes [7]. It has also been used to reveal the molecular channels in rhombohedral graphite structures, which can be used as an X-ray radiation filter [17]. We have also used sphere-in-contact models to model the complete reaction mechanism of heterogeneous catalytic reactions, where the adsorbates, the metal nanoparticle, and the substrate are explicitly shown in miniature models of heterogeneous catalysts [18]. When the sphere-in-contact model uses different radii for the atoms of the structure, then we can model many materials, either metallic, covalent, or ionic. Additionally, it has also been used to understand the position of nanoparticles on metal close-packed surfaces [18]. We have recently shown that sphere-in-contact models can be used to understand the number of (100) and (111) sites that a nanoparticle has on its exposed surface when such NPs are deposited on flat hexagonal, trigonal, and cubic surfaces of HCP and FCC lattices [19].
In this study, we show how the sphere-in-contact model can be used to model the repulsive adsorbate interactions during the adsorption of monoatomic elements (e.g., H, C, N, S, and O) on hexagonal metal surfaces of HCP and FCC lattices. The question we address with these physical molecular models is, ‘What is the thermodynamically most stable long-range ordering of monoatomic adsorbates on hexagonally close-packed surfaces, at a surface coverage of 1/3 ML?’. This is a question with great interest in surface chemistry, heterogeneous catalysis and nanotechnology and is addressed in this study using a pedagogic approach that connects math, physics and chemistry that is covered in the high-school curriculum to a scientifically more advanced research topic, such as the long-range ordering of surface adsorbates on metallic surfaces, that is studied at the university level surface chemistry and catalysis courses and relevant to interpreting experimental data from many surface analytical techniques, that are used in surface science to investigate such structures (e.g., AFM, STM, and LEED).
2. Materials and Methods
We have used marbles of different sizes and colors and a template that would hold the marbles in a hexagonal close-packed structure. The marbles are transparent so that we can observe the underlying structure beneath the adsorbates, such as the three-fold hollow sites in FCC and HCP lattices and the four-fold hollow sites in HCP lattices. We used a different color for the atoms in the metal surface and for the atoms in the adsorbate layer and a different radius for the spheres. Such a physical molecular model is not a close-packed model but rather a sphere-in-contact model, where the atoms have varied sizes and the surface density of adsorbates is less than expected for close-packed structures (e.g., FCC and HCP). The marbles have the natural tendency to arrange in a hexagonal close-packed pattern by lateral repulsive interactions, which can be enhanced by a lateral movement of the template frame. We have poured the 32 adsorbates onto the close-packed surface of the metal and observed where they would reside on the surface. All adsorbates resided in 3-fold hollow sites, and due to the relative radius of the metal and adsorbate atom, adsorption at the nearest neighbor 3-fold hollow site was not observed. We then moved the adsorbate atoms manually and randomly, one at a time, and observed how they would interact with other adjacent adsorbates. This process revealed collective adsorbate movement mechanisms on the surface of metals, such as the movement of an entire row of adsorbates to the next row of 3-fold hollow sites, due to repulsive interactions between the adsorbates, in a domino-type mechanism, when their electron cloud radius (i.e., atomic radius) was smaller than the radius of the metal electron cloud. So, the experimenter can study surface ordering phenomena using such physical models in a practical manner, which has pedagogical and research value. Additionally, we are able with this approach to study many possible surface ordering structures of adsorbates and obtain a picture of the structures that can be analyzed to calculate the repulsive energy between adsorbates and therefore suggest which long-range surface ordering is the thermodynamically more stable (i.e., lower in energy) structure.
This approach to studying surface adsorbate configurations on close-packed metallic surfaces and the quantitative analysis performed using trigonometry, geometry, and Coulomb’s law has not been previously published elsewhere to the best of our knowledge.
The adsorbate surface coverage (θ) was calculated by the assumption that we have full coverage (i.e., θ = 1, 1 ML) when, for each surface metal atom, there is one adsorbate atom available for surface chemical bonding. So, the equation of surface coverage used is
3. Results and Discussion
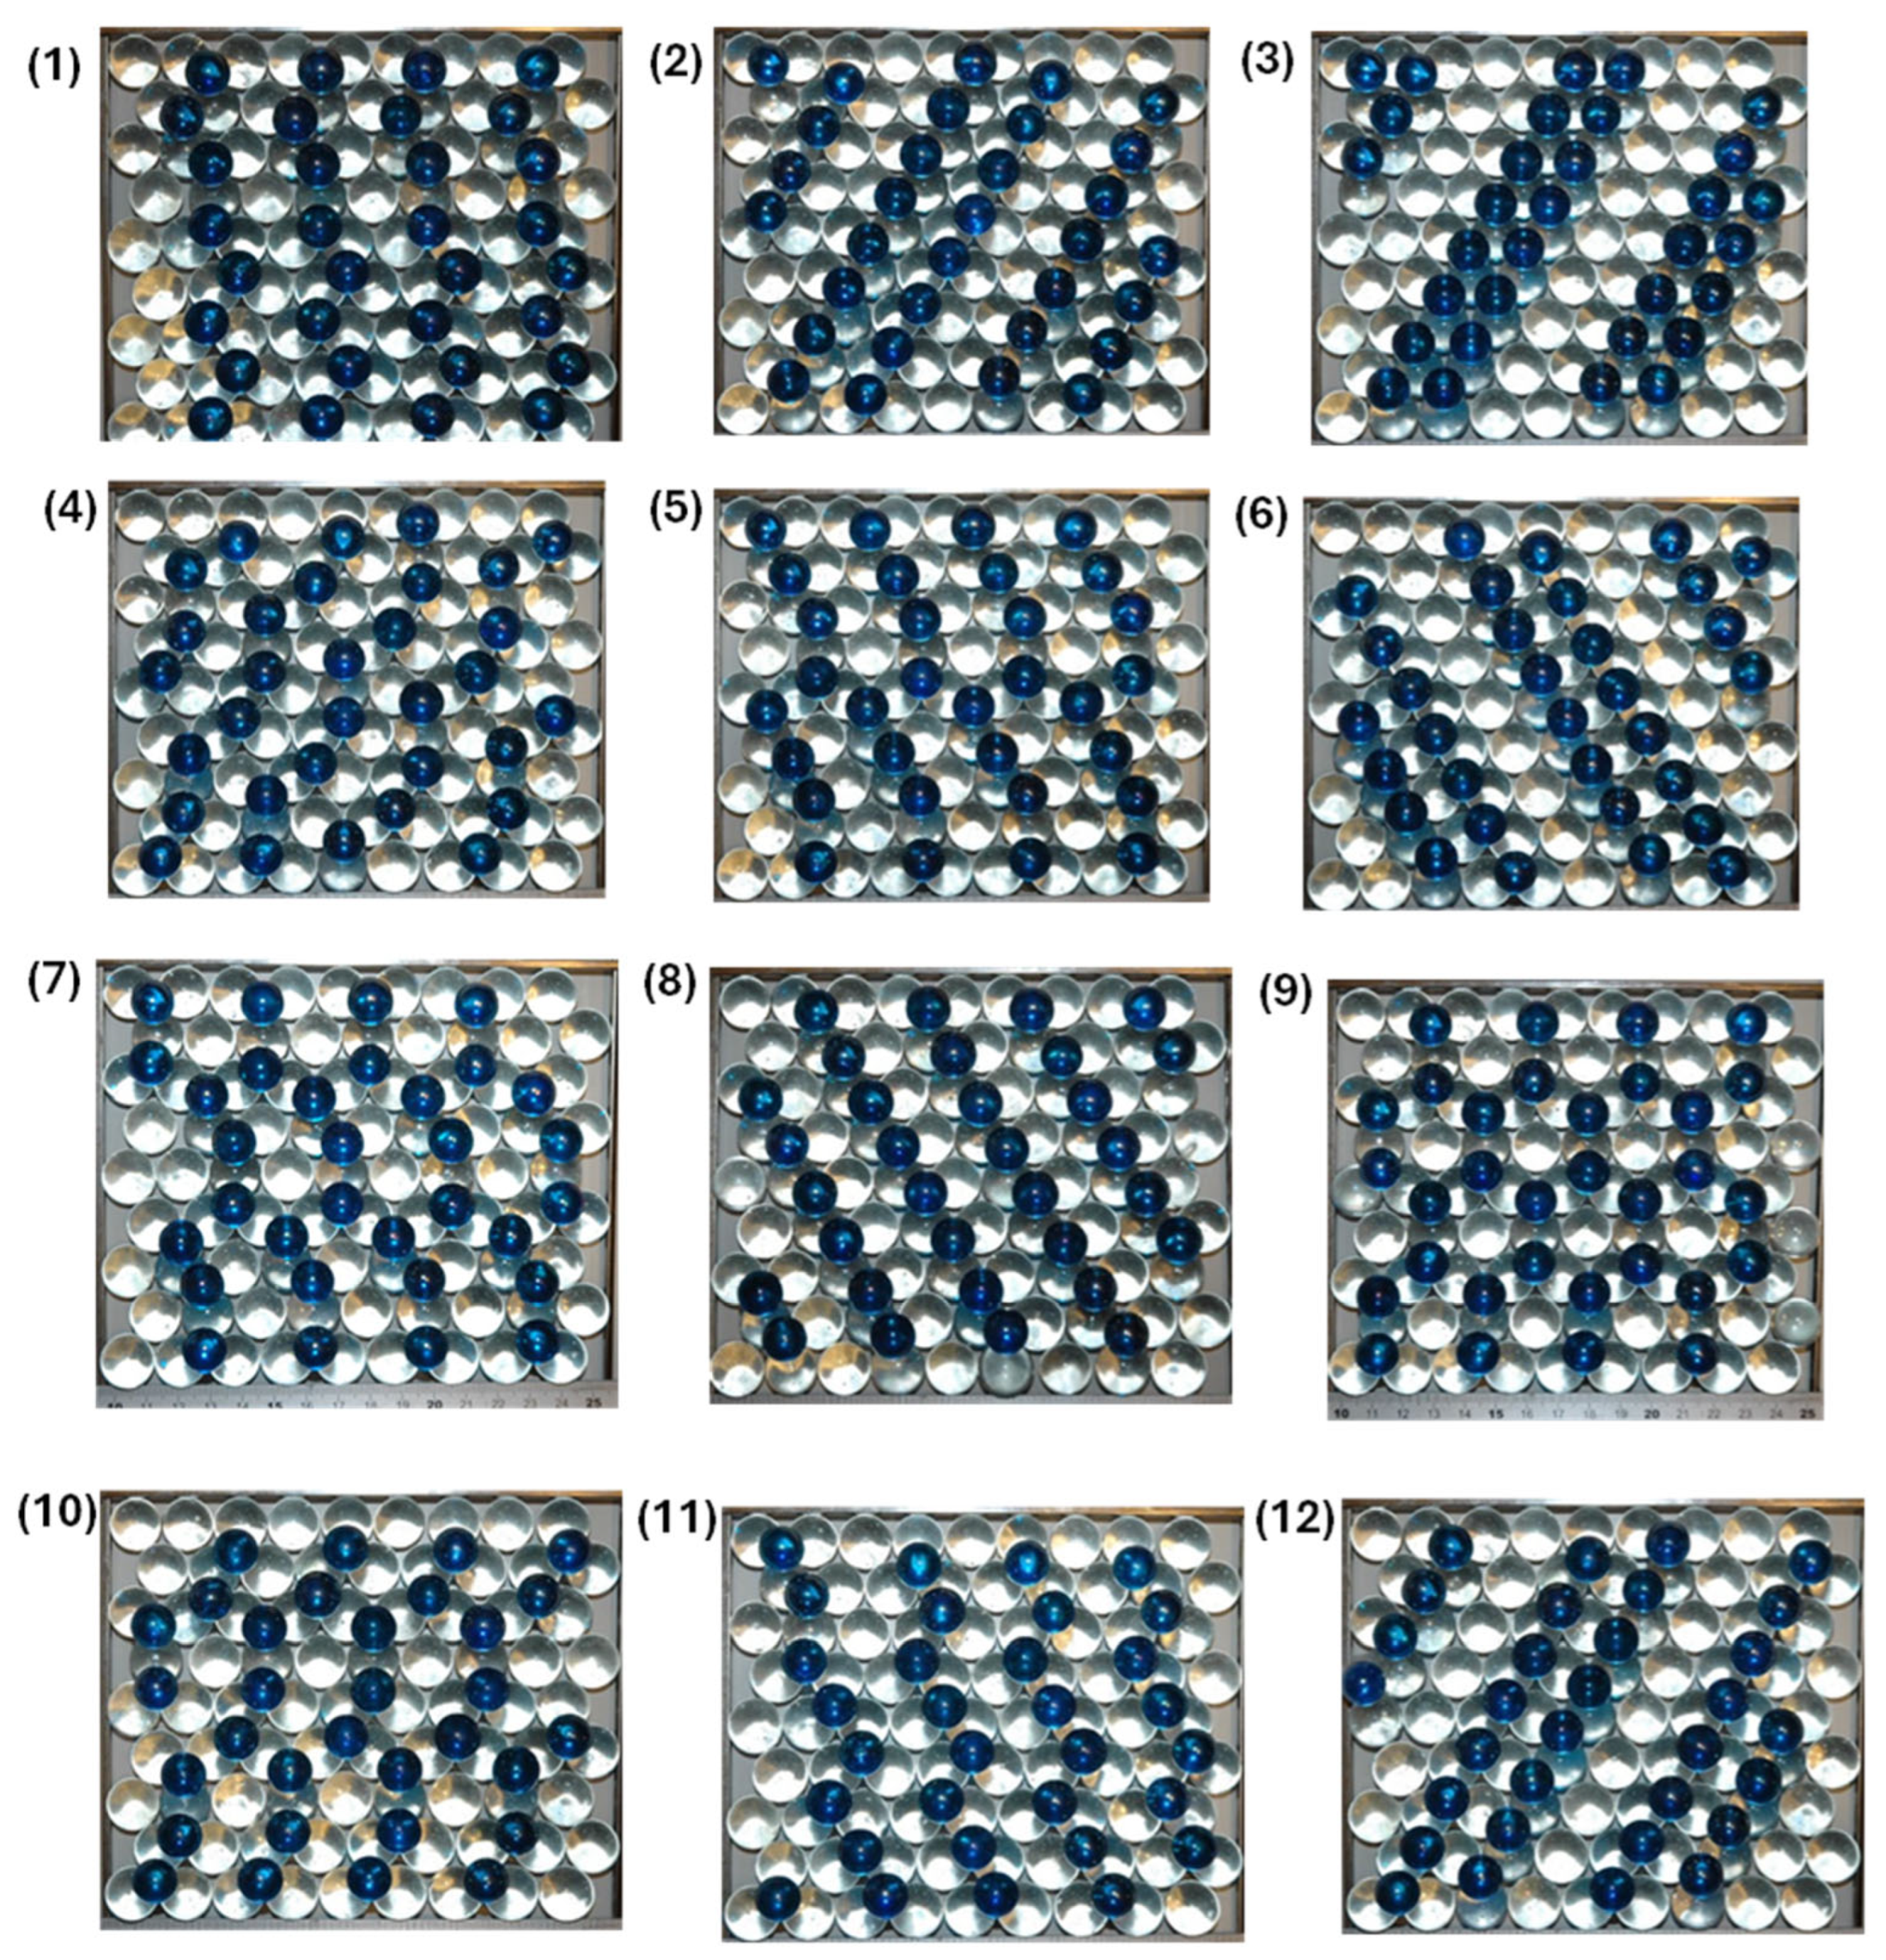
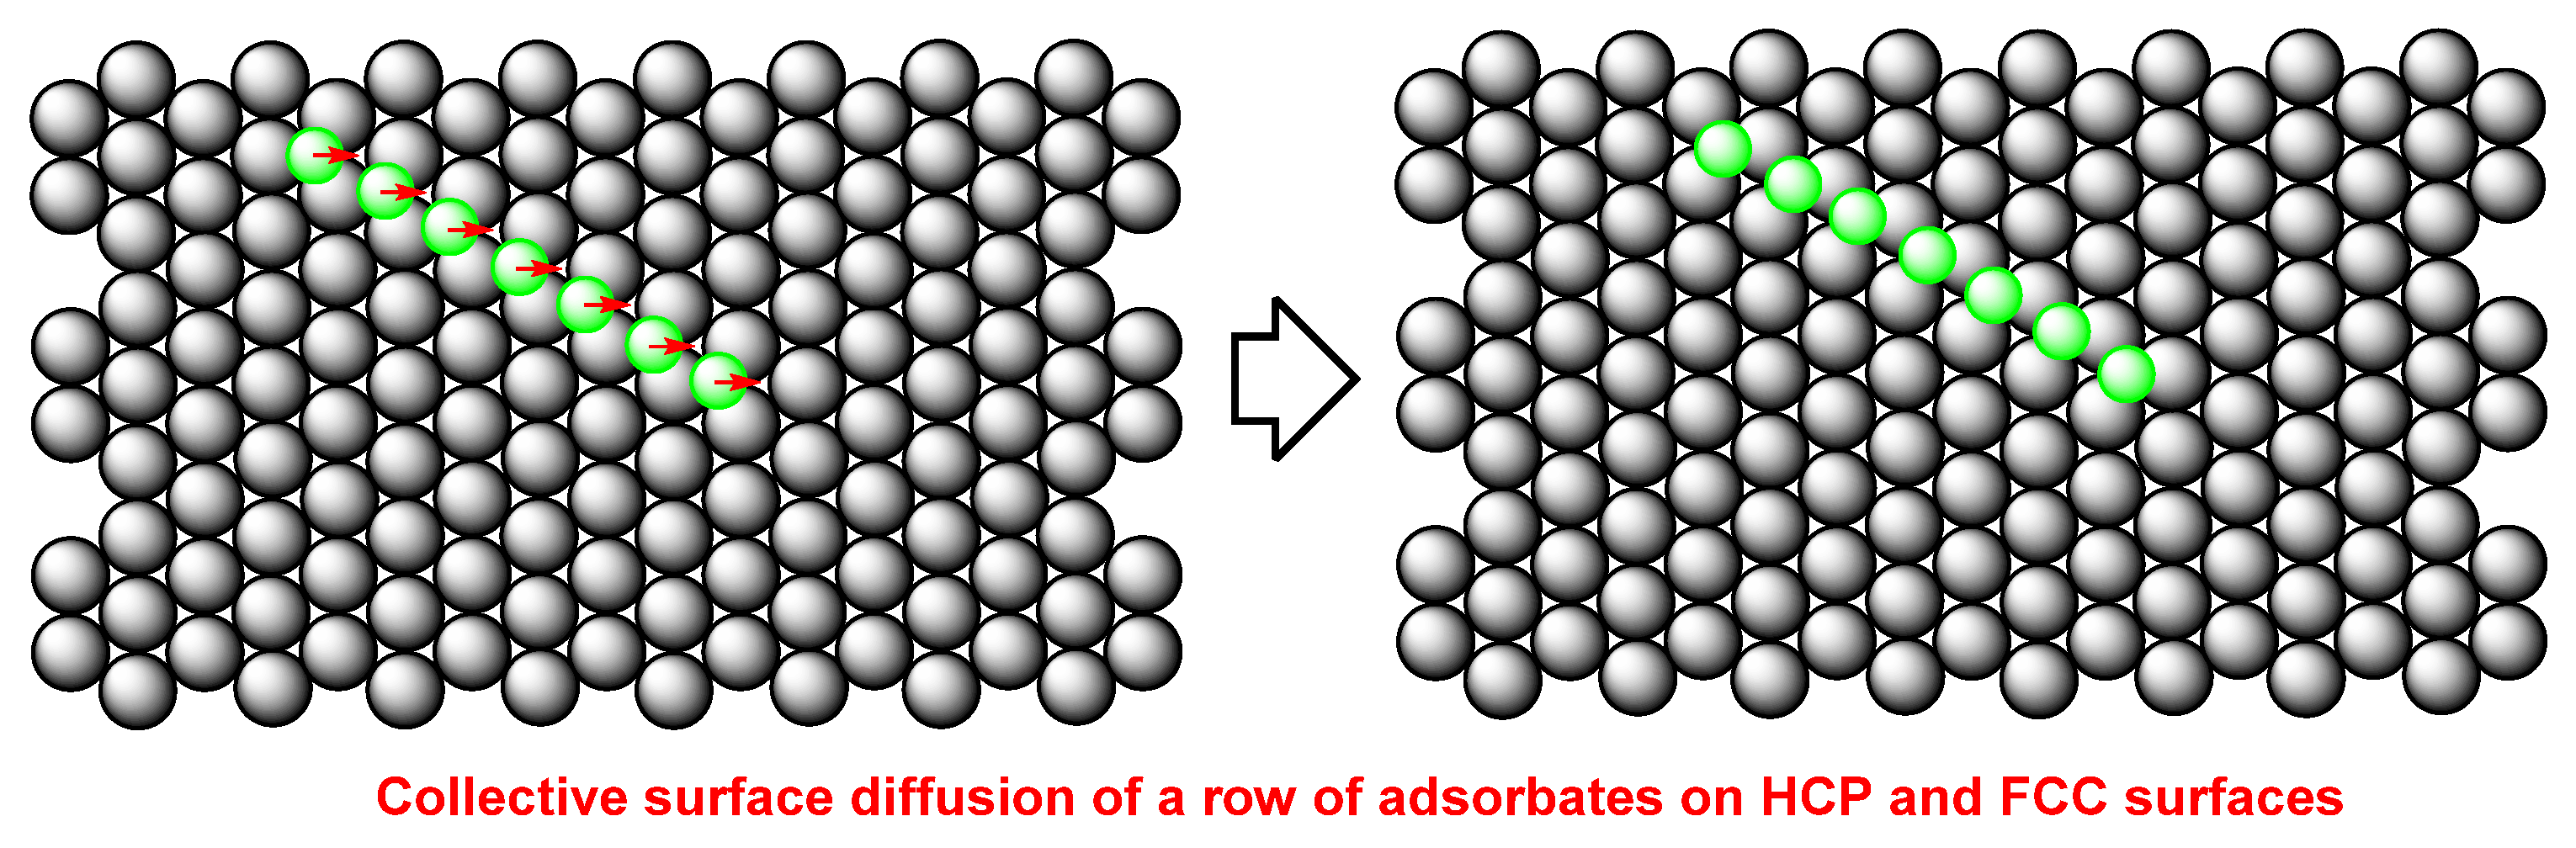
We have placed 81 colorless marbles in a rectangular template and allowed them to obtain their minimum energy configuration, which is a hexagonal close-packed structure. The sphere-in-contact models we have used are shown in Figure 1. All these physical molecular models have the same surface coverage, which is θ = 0.395 ML in the finite size of the supercell studied and corresponds to a surface coverage of 1/3 ML in the extended surface. This hexagonal close-packed structure is the surface termination of both hexagonal close-packed (HCP) and face-centered cubic (FCC) metals. However, similar models can be built for the body-centered cubic (BCC) structure. This means that all metallic surfaces, which usually have one of these three crystal structures, can be modeled using this approach, and surface adsorption can be studied in an interactive manner. Furthermore, steps and adatoms can be modeled within the sphere-in-contact model by adding a second layer of metal atoms or by adding an atom on the terrace of the metal surface, respectively. We have also prepared sphere-in-contact models with atom vacancies (e.g., defects), and such models could also be used to understand the mechanisms (i.e., the surface pathways) of surface adsorbate diffusion and saturation of surface defects in an interactive process. An example of a collective surface diffusion mechanism that was observed in these physical molecular models was the domino-like movement of surface adsorbates in a row, with adsorbates at the second nearest neighbor positions, to a row of adsorbate atoms displaced by one lateral lattice parameter (see Figure 2).
Figure 1.
Various ordered adsorptions of atoms on the surface of close-packed metals. Metal atoms are transparent, and the atoms of adsorbates are blue. The radius of the adsorbates is somewhat smaller than the radius of the metal atoms. We used 81 metal atom marbles and 32 adsorbate atom marbles to construct these images, where the adsorbate coverage was constant at θ = 0.395 ML. We note that in the infinite periodic cell, this would correspond to a surface coverage of 1/3 ML (0.333 ML).
Figure 2.
Collective surface diffusion of a row of adsorbate atoms (shown in green) to the nearest neighbor 3-fold hollow side through a domino-like surface diffusion mechanism on FCC and HCP surfaces.
Finally, such physical molecular models can also be used to model the adsorption of monoatomic adsorbates on hexagonally close-packed surfaces. Although such an approach only considers the geometric structure of adsorbates and does not include quantum effects on adsorption energies, it can be used as a starting point to evaluate different surface configurations of adsorbates, which could significantly reduce the computational cost of calculating the energy of adsorbate configurations in large unit cells explicitly. Therefore, we recommend the use of such physical sphere-in-contact models to understand dynamic processes in surface adsorption that are mainly a result of adsorbate repulsion and estimate the thermodynamic stability of adsorbate configurations of HCP and FCC metallic surfaces. Additionally, it can be readily used as a pedagogic method in the chemical sciences in fields such as catalysis, surface chemistry, and nanotechnology, as it offers an interactive tool to explore surface ordering phenomena of adsorbates on close-packed surfaces.
In some cases, a whole row of adsorbates can move simultaneously and form a new surface configuration in these physical sphere-in-contact models. These adsorbate–adsorbate interactions and the way the adsorbate atoms move on the surface to obtain new low-energy positions are an interactive research tool to explore potential collective diffusion mechanisms of adsorbates on surfaces, using an interactive and fast method of exploring many possible adsorbate surface configurations, various degrees of surface coverages, and the surface repulsion-induced rearrangement of adsorbates. This could suggest potential diffusion pathways that are currently unexplored and may aid in the interpretation of surface studies of adsorbates on metallic surfaces.
We note here that this model more suitable for adsorption studies where the adsorbates are in three-fold hollow sites, which are also the sites that usually have the highest adsorption energy (e.g., the most exothermic adsorption energy) compared to bridged and atop adsorption, which is also possible, especially when the adsorbate surface coverage increases and adsorbates are pushed due to repulsive interactions, to higher energy structures, that include both atop and bridged adsorption. We have previously studied the adsorption of CO on Pd nanoparticles and observed some of these trends in the adsorption energy of the 3-fold-hollow being generally greater than the bridged and top CO bound to the surface of the NP. If the crystal termination is due to a body-centered cubic (BCC) crystal, then the strongest adsorption site is usually the four-fold hollow site, followed by the bridged and the atop. These trends in adsorption energy we have previously shown are similar in large NPs that have well-defined surfaces and in small transition and noble metal clusters. However, the smaller clusters usually have more negative adsorption energies due to the unsaturated nature of the metal atoms on the cluster and the tendency to form stronger adsorbate-metal bonds [20].
It is therefore evident that this sphere-in-contact model is useful in understanding and presenting the adsorption of monoatomic adsorbates on the surface of metals. It can show the formation of various adsorbate adlayers with different adsorption configurations and long-range ordering of monoatomic adsorbates on metal surfaces. It can also help in the study of surface diffusion of adsorbates in a way that minimizes the repulsive interactions between adsorbates.
The fact that we use a different atomic radius for the adsorbates and the metal atoms makes it possible to measure the interatomic distances between the adsorbate atoms and calculate the repulsive interaction energy as a function of pairwise interactions using Coulomb’s law. These calculations follow in the subsequent section.
Analytical Equations of the Repulsive Interaction Energy of Monoatomic Adsorbates in the Sphere-in-Contact Model
Calculating the repulsive interaction energy in the sphere-in-contact model is possible by assuming that monoatomic adsorbates on the surface of a metal usually repel each other when they are adsorbed in nearby positions. Induced dipole—induced dipole interactions, such as London dispersion forces, will not be significant, as the adsorbate atoms need to be in physical contact to have the appearance of such electron cloud polarization effects. These repulsions are due to the valence electron cloud of adsorbates that is partially negatively charged, and since there is no attractive interatomic interaction between the adsorbates, such as dipole–dipole, ion–dipole, or ion–ion interactions, the only interaction left to evaluate is the Coulombic interaction between negatively charged partial charges on the surface of adsorbate atoms. This partial charge (δ) can be assumed to be smeared over the surface of a sphere with an atomic radius of the adsorbate atom. But to make the model of pairwise interactions even simpler, we can assume that the partial negative charge of the adsorbate atoms is at a point charge that is positioned at the nearest distance of the interacting adsorbate atom spheres. By making this assumption, we can calculate the pairwise repulsive interaction energy as a summation of Coulomb terms, that include all the surface adsorbates in a certain adsorption configuration. As this interaction energy has a 1/r dependence, it is fair to assume that it becomes relatively small at distances that exceed the fourth nearest neighbor. Therefore, in this analytical sphere-in-contact model, we include only pairwise interactions up to the third nearest neighbor. However, if these pairwise interactions are evaluated by a computer program, then the summation of pairwise interactions can be performed even for larger distances, which would result in a more accurate determination of the repulsive energy between adsorbates. Additionally, we model the adsorption at low surface coverages, and this suggests that the monoatomic adsorbates will bind to three-fold hollow sites. This has been observed for O and N adsorption on Ru(0001), where these species are adsorbed more exothermically at three-fold hollow sites rather than bridge and atop sites [20,21]. On Pt(111), the adsorption of atomic hydrogen has been found to be the most exothermic on the three-fold hollow site with ΔG = −0.333 eV at θ = 1/16 ML compared to ΔG = −0.285 eV at the bridge and ΔG = −0.255 eV at the atop sites [22]. Density Functional Theory (DFT) calculations have shown that H species adsorb more exothermically on Ni, Pd, and Pt (111) surfaces at three-fold hollow sites and that the difference between three-fold hollow site adsorption and bridge adsorption can be used as an indicator of the barrier of surface diffusion [23]. Additionally, a recent dispersion-corrected DFT study on 20 transition metals (TMs) close-packed surfaces (i.e., FCC(111) and HCP(0001)) showed that H adsorption is more favorable at 3-fold hollow sites for FCC metals (e.g., Ag, Au, Cu, Ir, Ni, Pd, Pt, and Rh) and HCP metals (e.g., Co, Hf, Os, Re, Ru, Sc, Tc, Ti, Y, and Zr) with the only exception of Cd and Zr, where bridge adsorption was favorable [24]. Three-fold hollow adsorption at low coverages has also been observed for diatomic molecules such as CO, where the adsorption occurs primarily through an end-on configuration on Pd NPs [25]. Finally, there is STM evidence that N adsorption on Ru(0001) surfaces is more stable at 3-fold hollow sites [26]. Therefore, it is reasonable to assume that on metallic close-packed HCP and FCC surfaces, the adsorption of adsorbates will start on three-fold hollow sites at low coverages and then move to bridge and atop sites as the coverage increases, and adsorbate-adsorbate and adsorbate-substrate electronic effects become more important.
Coulomb’s law for the force between interacting particles is
where F is the electric force (units = N), k is the Coulomb constant (8.988 × 10⁹ Nm2/C2), and are the point charges on the adsorbate surface and r the distance of separation between the adsorbate surfaces in m. As we will model the repulsive interaction between the adsorbates using a partial charge located at the nearest distance points of the adsorbate spheres, this energy is more exothermic when the particle separation is small. The interaction energy (U) of Coulomb’s law is given by
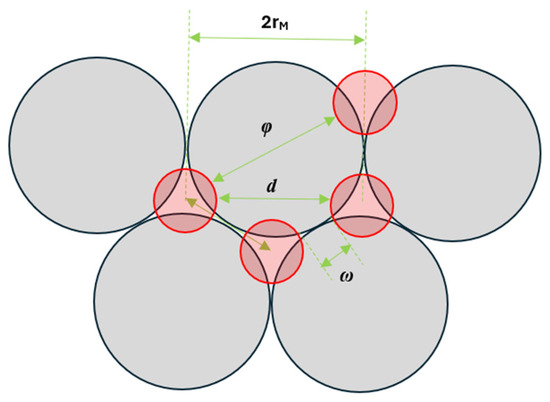
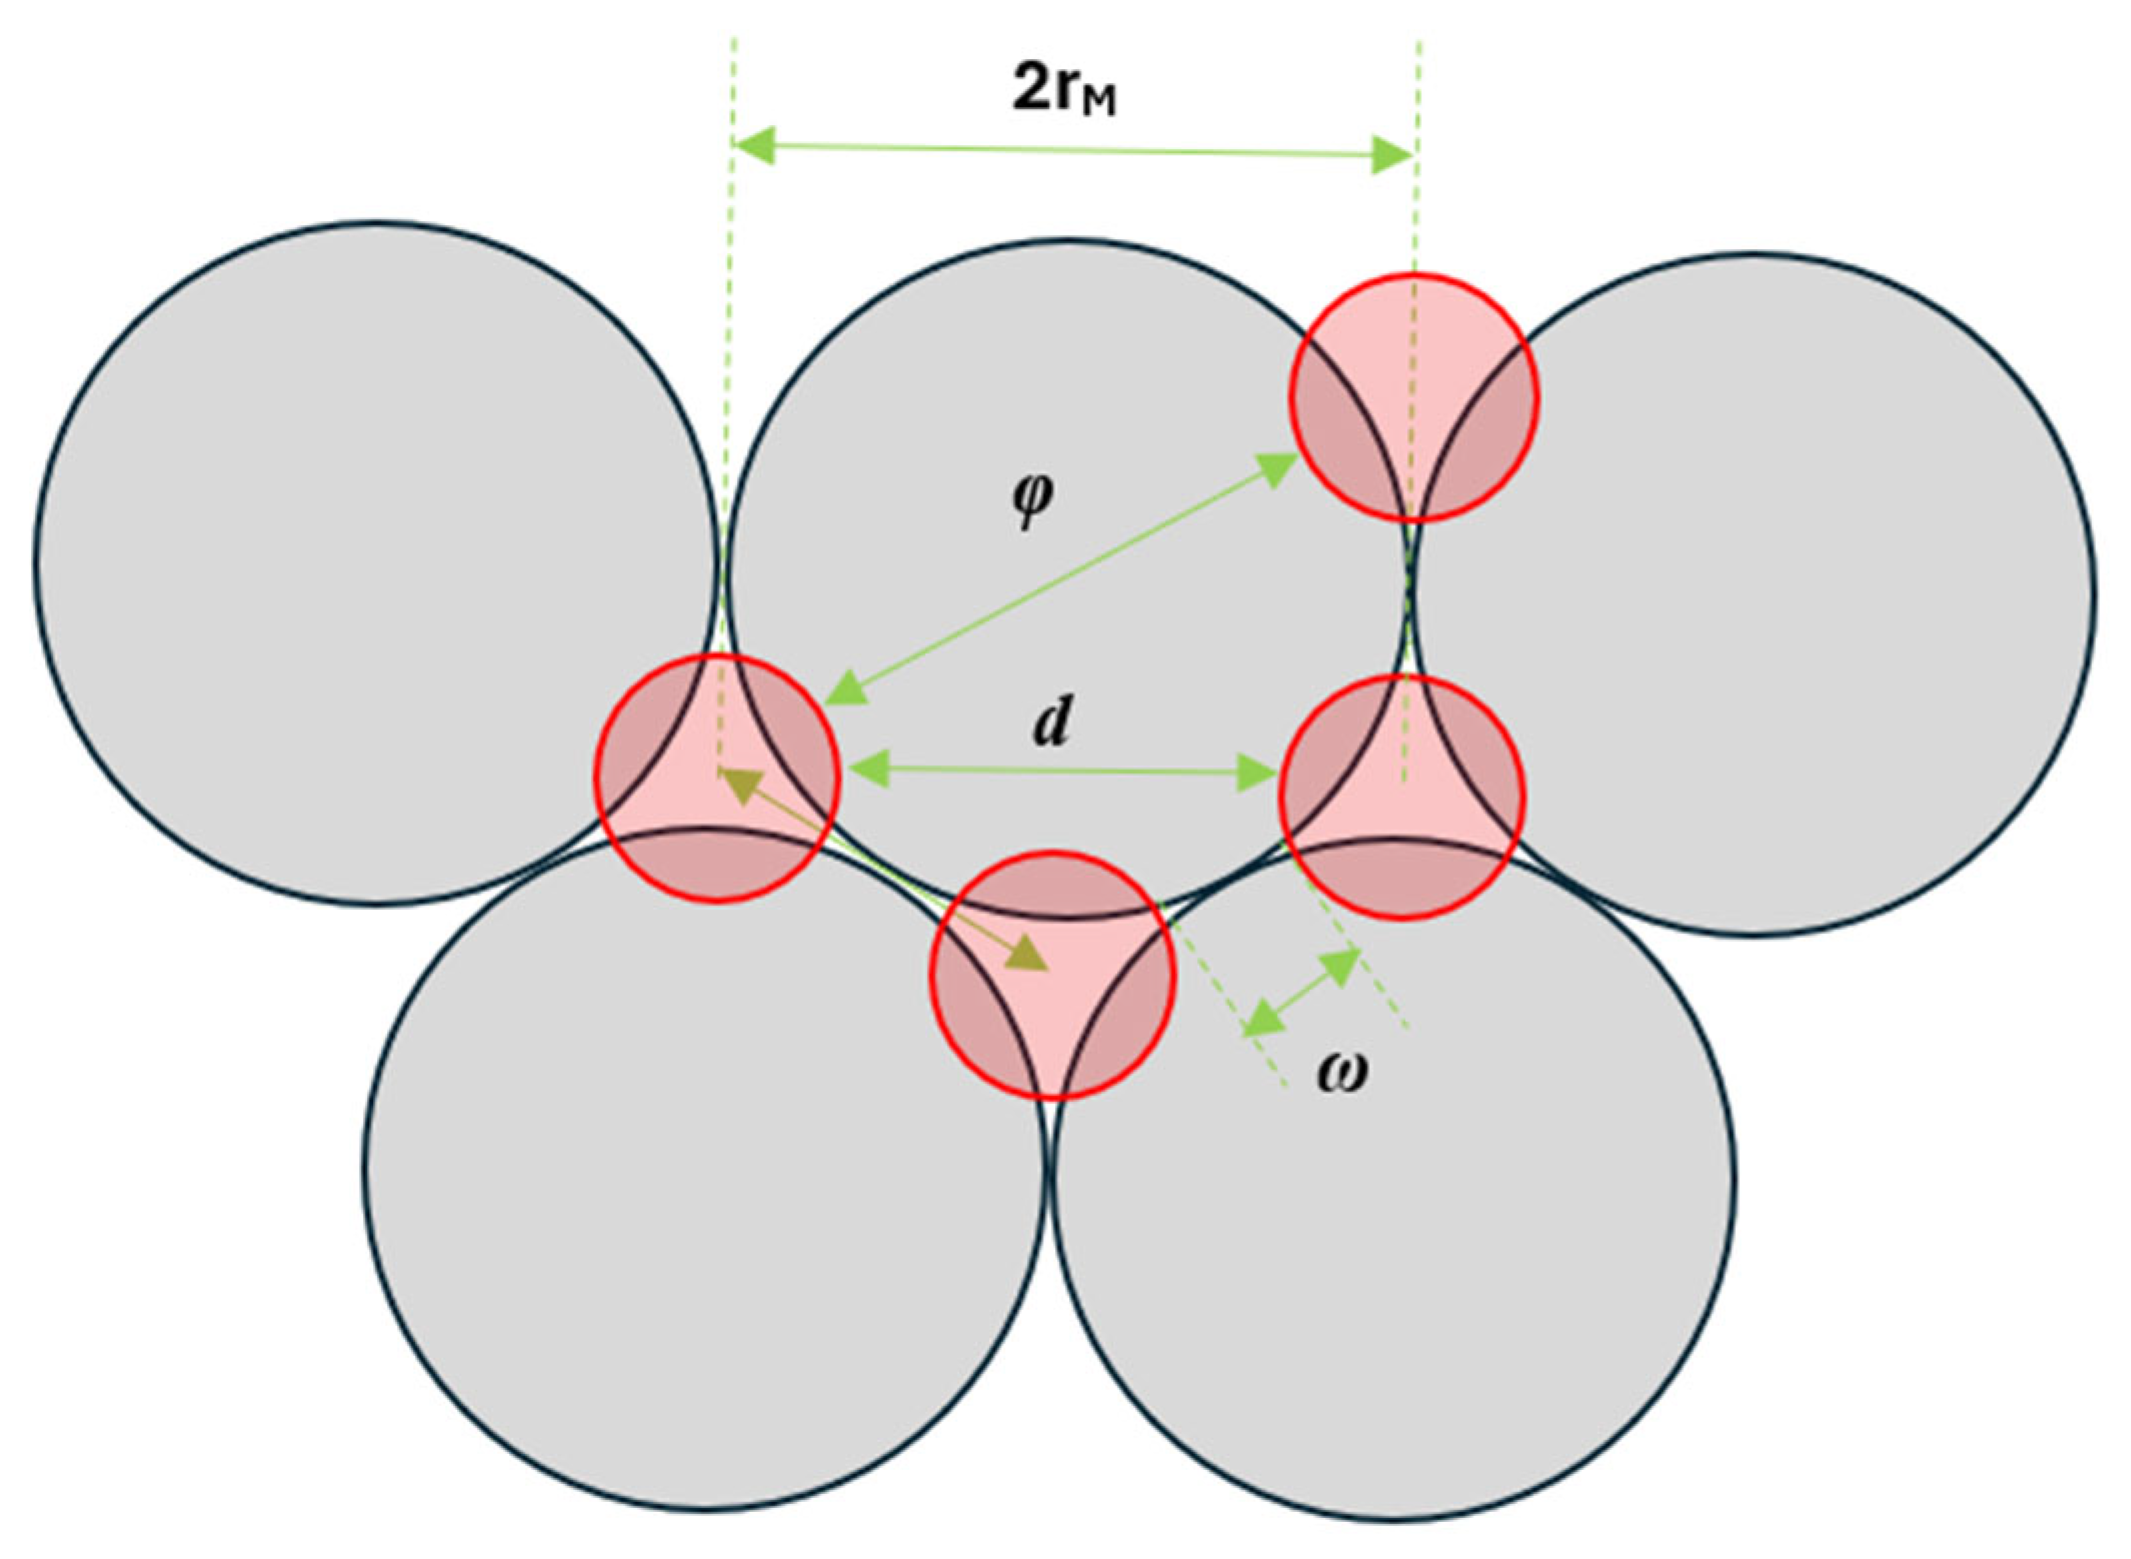
The pairwise interactions will be modeled this way with a summation of all interacting adsorbate particles that are in nearest neighbor, second nearest neighbor, and third nearest neighbor three-fold hollow sites. When the point charges are assumed to be on the surface of the adsorbate spheres, at points that are defined by a line being the nearest distance between the adsorbate spheres, then the distance between the nearest neighbors is ω, d, and φ, as shown in Figure 3.
Figure 3.
Monoatomic adsorbate atoms are shown by red circles, adsorbed on the close-packed surface of metal atoms, shown by black circles. The distances ω, d, and φ are the nearest neighbor, second nearest neighbor, and third nearest neighbor distances between adsorbate atoms at three-fold hollow sites, respectively.
The energy of the repulsive Coulomb interaction energy will be proportional to how close the adsorbate spheres (i.e., atoms) are when they are in 3-fold hollow sites. This provides the possibility of modeling surface adsorption on large surface unit cells with various adsorption configurations. These sphere-in-contact models can exhibit long-range ordering phenomena, as shown in the configurations of adsorbates in Figure 1.
Therefore, the net repulsive Coulomb interaction energy is given by the summation of first, second, and third nearest neighbor interactions between adsorbates adsorbed at the three-fold hollow sites of close-packed FCC and HCP metal surfaces, given by
The nearest distance between adsorbates in the second closest 3-fold hollow positions is given by
where is the radius of the metal atoms and the radius of the adsorbate which is depicted in Figure 3.
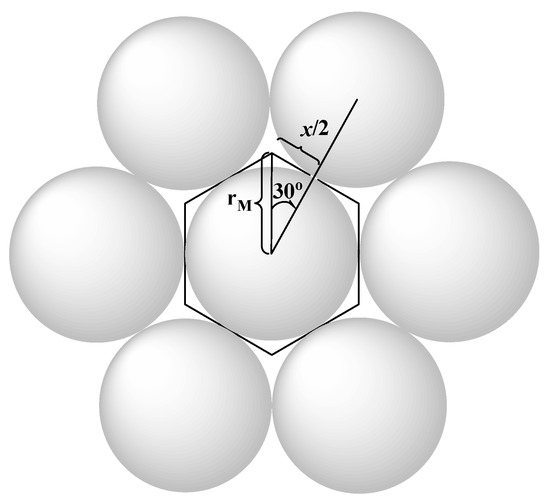
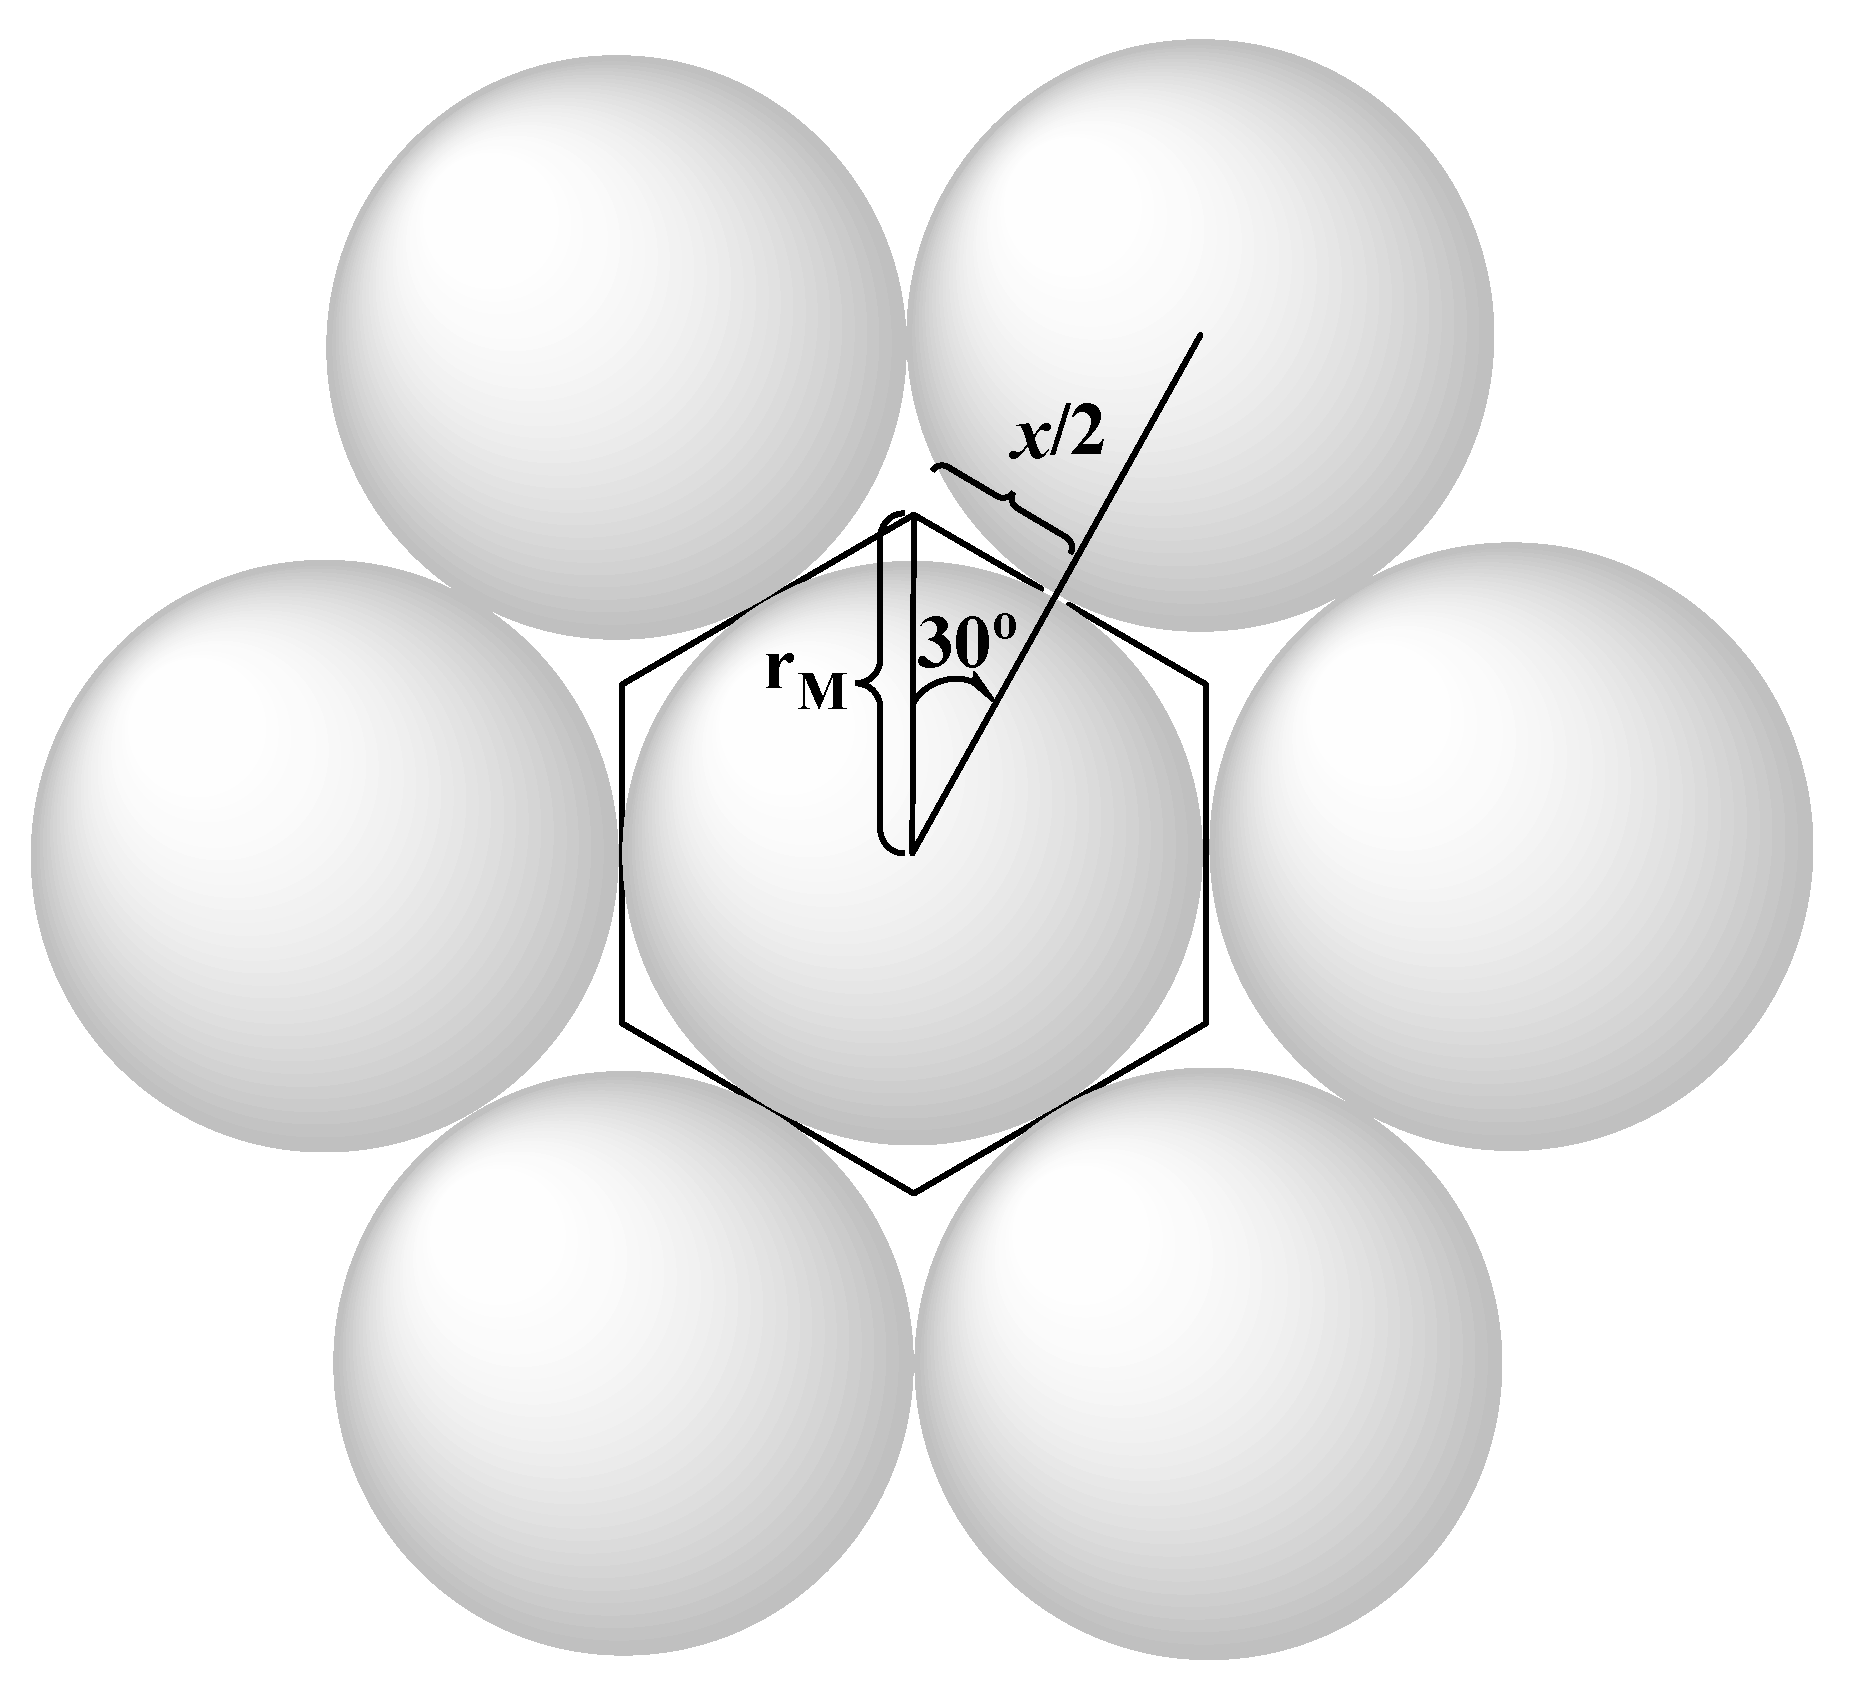
The centers of the adsorbate spheres in nearest neighboring sites have a distance of x. If you consider the six nearest neighbor distances that surround each metal atom, then these form a regular hexagon shown in Figure 4.
Figure 4.
Simplified schematic that presents the regular hexagon formed by nearest neighbor sites on closed-packed metal surfaces of HCP and FCC lattices.
The side of this hexagon is x, and the distance from the center of the hexagon to the side of the hexagon is the radius of the metal atom, . If you also consider the radius of the hexagon, then this, along with x/2 and , form a 30-60-90 triangle from which you can calculate x from the tangent of the 30° angle. This yields that
So, the nearest neighbor distance between the surfaces of the spheres of adsorbates at 3-fold hollow sites of HCP and FCC crystals is given by
Therefore, combining Equations (6) and (7) yields that the nearest neighbor distance between adsorbates at three-fold hollows is
The third nearest neighbor distance between adsorbates is given by the long diagonal of the regular hexagon in Figure 3, the length of which can be calculated using the Pythagorean theorem. So, the distance between the point charges of the third nearest neighbor adsorbates is given by
This means that the relative pairwise repulsive interaction energy between adsorbates given from Equations (4), (5), (8), and (9) is
In Table 1, we evaluate the repulsive interaction energy of the various sphere-in-contact models shown in Figure 1 using Equation (10). In the sphere-in-contact models of Figure 1, the adsorbates have a radius and therefore nearest neighbor adsorption is not observed in any of the models. However, second-nearest-neighbor and third-nearest-neighbor absorption is observed. The number of pairwise interactions for second (v2) and third (v3) nearest neighbor interactions is given in Table 1, along with the repulsive interaction energy (Unet) per 1 mol of adsorbates for adsorbates having partial charges of −0.1 e and −0.2 e. We note here that in this model, where only repulsive interactions between adsorbate atoms and their geometric inter-surface distance are considered, the relative value and sign of the partial charge do not affect the trend in stabilities of these adsorbate arrangements at low surface coverages.
Table 1.
Showing the various repulsive adsorption energies for the complete adsorbate surface sphere-in-contact model and the number of nearest neighbor interactions that were found in each different surface configuration shown in Figure 1. The adsorbate partial charge was either −0.1 e or −0.2 e.
As the coverage used in our sphere-in-contact models was θ = 0.395 ΜL, therefore we only consider adsorption in the three-fold hollow sites. Here we note that currently surface adsorption in physical sphere-in-contact models is currently only possible at 3-fold hollows and 4-fold hollows; however, it is conceivable that such physical molecule models using magnetic spheres can potentially also result in bridge and atop adsorption. A surface coverage of 0.395 ML was chosen, primarily because adsorption on hexagonally close-packed metallic surfaces is expected to be thermodynamically more stable at the 3-fold hollow sites at low surface coverages, and at higher coverages, adsorption would also populate the bridge and atop adsorption sites that have a smaller number of chemical bonds to the adsorbate.
Then, by using the radius of the metal atom of = 1.5 Å, the adsorbate = 1.3 Å, and a partial negative charge of −0.1 e and −0.2 e for and , we can calculate using Equation (10) the repulsive interaction energy of different surface configurations of adsorbates of the 12 adsorption configurations shown in Figure 1. The partial charge (δ) of each adsorbate is usually negative due to charge transfer from the metal to the adsorbatewhich results in a repulsive energy interaction between adsorbates. These analytical calculations reveal that surface 9 has the lowest energy configuration for monoatomic adsorbates at a θ of 1/3 ML. This is a surface configuration of the adsorbates with hexagonal symmetry in which all adsorbates are arranged at the third nearest neighbor adsorption sites. We suggest that this adsorption configuration should be sought for in surface imaging techniques (e.g., STM and AFM) and surface structure determination techniques (e.g., LEED) in order to proof that long-range ordering of monoatomic adsorbates on close-packed HCP and FCC metal surfaces is dictated by repulsive surface interactions between adsorbate atoms, which are minimized at the third nearest neighbor 3-fold hollow sites at coverages of 1/3 ML, when thermodynamic annealing occurs that allows the diffusion of adsorbates to low energy surface configurations. Structures of this symmetry have been observed at a coverage of 1/3 for sulfur adsorption on Au(111), which are, however, at the 3rd nearest neighbor sites, possibly due to lower surface coverage [27].
The detailed understanding of surface adsorption of monoatomic adsorbates and adsorbates bound through a single atom on the surface of metals is of fundamental interest to surface science, materials science, and heterogeneous catalysis. The simplified approach followed in this manuscript can capture the effect of lateral repulsions between any monoatomic adsorbate bound at 3-fold hollow sites and determine the surface adsorption configuration that is the lowest in repulsive interaction energy. This approach can also be used to study the coverage-dependent adsorption energy trends of adsorbates as a function of repulsive lateral electrostatic interactions [28], which have been suggested to affect the adsorption energy and desorption barrier of hydrogen on metals.
4. Conclusions
We have used the sphere-in-contact model to study the structure of monoatomic adsorbates on hexagonal close-packed surfaces of (0001)-HCP and (111)-FCC metals. This model contains atoms of different radii and colors to represent the atoms of the metal surface and the adsorbate atoms. The sphere-in-contact model allows the formation of adsorbate adlayers of different adsorption configurations in an interactive way, exploring many possible surface configurations and evaluating which has the lowest energy using analytical trigonometric equations and well-established equations in physics.
The study reveals that using this approach, we can get the arrangement of the adsorbates on metal surfaces at various coverages and compare structures of different surface adsorbate configurations that were obtained for the same surface coverage. This has useful applications in an educational context but could also be used as a research method to determine the lowest energy adsorption configuration of adsorbates when repulsive lateral interactions are important.
With the use of this approach, we find that at a surface adsorbate coverage of θ = 1/3 ML, monoatomic adsorbates will form a hexagonal surface adlayer in which the adsorbates are at the 3rd nearest neighbor 3-fold hollow sites. This can have useful implications in the use of such adlayers in technological applications such as coatings and arrays of magnetic atoms on surfaces. Furthermore, we anticipate that this approach will help teachers, researchers, and students to visualize the structure of adsorbates on metal nanoparticles and surfaces in a practical way that allows real-time manipulation and visualization of repulsive adsorbate–adsorbate interactions and the observation of collective adsorbate diffusion mechanisms due to lateral adsorbate repulsions. It also allows the study of the energetic stability of disordered H adsorption, as this has been suggested on the surface of Ni(111) through Low Energy Electron Diffraction (LEED) experiments [29].
Funding
This research received no external or internal funding.
Data Availability Statement
The original contributions presented in this study are included in the article. Further inquiries can be directed to the corresponding author.
Acknowledgments
The author acknowledges Surfaces for covering the APC charge of the manuscript to make it available as open access.
Conflicts of Interest
There are no conflicts of interest to declare in this study.
Abbreviations
The following abbreviations are used in this manuscript:
| ML | Monolayer |
| HCP | Hexagonal close-packed |
| FCC | Face-centered cubic |
| BCC | Body-centered cubic |
| DFT | Density functional theory |
| STM | Scanning tunneling microscopy |
| AFM | Atomic force microscopy |
| LEED | Low-energy electron diffraction |
| NP | Nanoparticle |
| CPK | Corey–Pauling–Koltun |
| VdW | Van der Waals |
References
- Wang, Z.; Hu, P. Rational catalyst design for CO oxidation: A gradient-based optimization strategy. Catal. Sci. Technol. 2021, 11, 2604–2615. [Google Scholar] [CrossRef] [Scilit]
- Xu, H.; Cheng, D.; Cao, D.; Zeng, X.C. Revisiting the universal principle for the rational design of single-atom electrocatalysts. Nat. Catal. 2024, 7, 207–218. [Google Scholar] [CrossRef] [Scilit]
- Saini, S.; Halldin Stenlid, J.; Abild-Pedersen, F. Electronic structure factors and the importance of adsorbate effects in chemisorp-tion on surface alloys. npj Comput. Mater. 2022, 8, 163. [Google Scholar] [CrossRef] [Scilit]
- Sprunger, P.T.; Plummer, E.W. The Interaction of Hydrogen with Simple Metal Surfaces. Surf. Sci. 1994, 307–309, 118–123. [Google Scholar] [CrossRef] [Scilit]
- Comer, B.M.; Li, J.; Abild-Pedersen, F.; Bajdich, M.; Winther, K.T. Unraveling Electronic Trends in O* and OH* Surface Adsorption in the MO2 Transition-Metal Oxide Series. J. Phys. Chem. C 2022, 126, 7903–7909. [Google Scholar] [CrossRef] [Scilit]
- Raman, A.S.; Vojvodic, A. Energy Trends in Adsorption at Surfaces; Springer International Publishing: Cham, Switzerland, 2020; pp. 1321–1341. [Google Scholar] [CrossRef] [Scilit]
- Zeinalipour-Yazdi, C.D.; Loizidou, E.Z. Study of the cap structure of (3,3), (4,4) and (5,5)-SWCNTs: Application of the sphere-in-contact model. Carbon 2017, 115, 819–827. [Google Scholar] [CrossRef] [Scilit]
- Extremera, J.; Vergara, D.; Rodríguez, S.; Dávila, L.P. Reality-Virtuality Technologies in the Field of Materials Science and Engineering. Appl. Sci. 2022, 12, 4968. [Google Scholar] [CrossRef] [Scilit]
- Zeinalipour-Yazdi, C.D.; Pullman, D.P.; Catlow, C.R.A. The sphere-in-contact model of carbon materials. J. Mol. Model. 2016, 22, 40. [Google Scholar] [CrossRef] [Scilit]
- Corey, R.B.; Pauling, L. Molecular models of amino acids, peptides, and proteins. Rev. Sci. Instrum. 1953, 24, 621–627. [Google Scholar] [CrossRef] [Scilit]
- Zeinalipour-Yazdi, C.D. Electronic Structure and Interlayer Binding Energy of Graphite; University of California: San Diego, CA, USA, 2006. [Google Scholar]
- Zeinalipour-Yazdi, C.D.; Pullman, D.P. Correlation of Polarizabilities with Van Der Waals Interactions in π-systems. J. Phys. Chem. B 2006, 110, 24260–24265. [Google Scholar] [CrossRef] [Scilit]
- Wert, C.A. Diffusion coefficient of C in α-iron. Phys. Rev. 1950, 79, 601–605. [Google Scholar] [CrossRef] [Scilit]
- Ågren, J. A revised expression for the diffusivity of carbon in binary Fe-C austenite. Scr. Metall. 1986, 20, 1507–1510. [Google Scholar] [CrossRef] [Scilit]
- Goldschmidt, H.J. Metal Physics of Interstitial Alloys. In Interstitial Alloys; Springer: Boston, MA, USA, 1967; Chapter 2; pp. 37–59. [Google Scholar] [CrossRef] [Scilit]
- Fukai, Y. Site preference of interstitial hydrogen in metals. J. Less Common Met. 1984, 101, 1–16. [Google Scholar] [CrossRef] [Scilit]
- Zeinalipour-Yazdi, C.D.; Pullman, D.P. Study of a rhombohedral graphite X-ray filter using the sphere-in-contact model. Chem. Phys. Lett. 2019, 734, 136717. [Google Scholar] [CrossRef] [Scilit]
- Zeinalipour-Yazdi, C.D.; Pullman, D.P. Miniature physical sphere-in-contact models of heterogeneous catalysts and metal nanoparticles. J. Mol. Model. 2023, 29, 307. [Google Scholar] [CrossRef] [Scilit]
- Zeinalipour-Yazdi, C.D. A study using physical sphere-in-contact models to investigate the structure of close-packed nanoparticles supported on flat hexagonal, square and trigonal lattices. Chem. Phys. 2025, 588, 112464. [Google Scholar] [CrossRef] [Scilit]
- Kostov, K.L.; Gsell, M.; Jakob, P.; Moritz, T.; Widdra, W.; Menzel, D. Observation of a novel high density 3O(2 × 2) structure on Ru(001). Surf. Sci. 1997, 394, L138–L144. [Google Scholar] [CrossRef] [Scilit]
- Mortensen, J.J.; Hammer, B.; Nørskov, J.K. A theoretical study of adsorbate-adsorbate interactions on Ru(0001). Surf. Sci. 1998, 414, 315–329. [Google Scholar] [CrossRef] [Scilit]
- Liu, J.; Hagopian, A.; McCrum, I.T.; Doblhoff-Dier, K.; Koper, M.T.M. Unraveling the Origin of the Repulsive Interaction between Hydrogen Adsorbates on Platinum Single-Crystal Electrodes. J. Phys. Chem. C 2024, 128, 15019–15028. [Google Scholar] [CrossRef] [Scilit]
- Watson, G.W.; Wells, R.P.K.; Willock, D.J.; Hutchings, G.J. A Comparison of the Adsorption and Diffusion of Hydrogen on the {111} Surfaces of Ni, Pd, and Pt from Density Functional Theory Calculations. J. Phys. Chem. B 2001, 105, 4889–4894. [Google Scholar] [CrossRef] [Scilit]
- Allés, M.; Meng, L.; Beltrán, I.; Fernández, F.; Viñes, F. Atomic Hydrogen Interaction with Transition Metal Surfaces: A High-Throughput Computational Study. J. Phys. Chem. C 2024, 128, 20129–20139. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Zeinalipour-Yazdi, C.D.; Willock, D.J.; Thomas, L.; Wilson, K.; Lee, A.F. CO adsorption over Pd nanoparticles: A general framework for IR simulations on nanoparticles. Surf. Sci. 2016, 646, 210–220. [Google Scholar] [CrossRef] [Scilit]
- Trost, J.; Zambelli, T.; Wintterlin, J.; Ertl, G. Adsorbate-adsorbate interactions from statistical analysis of STM images: N/Ru(0001). Phys. Rev. B 1996, 54, 17850. [Google Scholar] [CrossRef] [Scilit]
- Abufager, P.N.; Zampieri, G.; Reuter, K.; Martiarena, M.L.; Busnengo, H.F. Long-Range Periodicity of S/Au(111) Structures at Low and Intermediate Coverages. J. Phys. Chem. C 2014, 118, 290–297. [Google Scholar] [CrossRef] [Scilit]
- Xu, L.; Ma, Y.; Zhang, Y.; Teng, B.; Jiang, Z.; Huang, W. Revisiting H/Pt(111) by a combined experimental study of the H-D exchange reaction and first-principles calculations. Sci. China Chem. 2011, 54, 745–755. [Google Scholar] [CrossRef] [Scilit]
- Christmann, K.; Schober, O.; Ertl, G.; Neumann, M. Adsorption of hydrogen on nickel single crystal surfaces. J. Chem. Phys. 1974, 60, 4528–4540. [Google Scholar] [CrossRef] [Scilit]

Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2026 by the author. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license.