


Abstract
Potato (Solanum tuberosum L.) is a vital food security and cash crop in Eritrea, yet bacterial and fungal pathogens responsible for 15–30% yield losses remain molecularly uncharacterized across its production systems. Here, we present the first nationwide amplicon-based metagenomic survey of potato pathogen communities, sampling 81 farms across 14 sub-regions spanning four agroclimatic regions during July–August 2023. High-throughput amplicon sequencing targeting the bacterial 16S rRNA V3–V4 region and fungal ITS1–ITS2 loci revealed pronounced geographic heterogeneity in community composition and alpha diversity. Pseudomonas spp. were ubiquitous, with a peak relative abundance of 32.5% in Dekemhare. Dominant fungi included Alternaria spp. (14.3% in Berik), Fusarium spp. (highest diversity 53.8% in Adi Kuala), Botrytis cinerea (36.9% in Adi Keih), and Rhizoctonia solani (44.4% in Adi Tekeliezan). Bacterial Shannon diversity averaged 5.67; fungal, 4.70. Weighted UniFrac PCoA accounted for 56.5% of bacterial community variance along PC1, confirming distinct geographic clustering. Potential pathogen-associated taxa of Pseudomonas, Alternaria, Fusarium, and Colletotrichum were detected in every asymptomatic sample examined, demonstrating the inadequacy of visual-only disease surveillance. These findings establish the first molecular baseline for potato-associated taxon diversity in Eritrea, providing the empirical foundation for region-specific integrated disease management and evidence-based seed certification protocols.
1. Introduction
Potato (Solanum tuberosum L.) is an important food and cash crop supporting many populations around the world for both income generation and ensuring food security. However, its production is seriously challenged by several biotic and abiotic stresses, where bacterial and fungal pathogens are the most economically important pathogens, leading to severe yield reductions up to 15–30% in East African countries [1]. In Eritrea, potatoes are mainly grown in the highlands of Maekel and Debub regions. The medium climatic conditions in these regions provide the optimal conditions for crop growth, while the crops’ nature facilitates colonization by pathogens that lead to disease and spread from generation to generation. Understanding and managing disease requires full information on the causative agents’ biology, genetic variation, and the spatial distribution patterns in the specific production environments. Conventional disease surveillance methods have been heavily dependent on visual symptom identification and isolation by culture-based techniques [2]. However, such conventional methods can only give a pathogen profile of the already reported pathogen diversity in the region using target primers or reference materials. This creates incomplete information for pathogens that show similar symptoms, and pathogens that are not easily grown on culture media are difficult to identify. Moreover, the absence of visible symptoms, such as the emerging recombinant strains of Potato Virus Y (PVY) [3], often leads to the omission or underestimation of important yield-reducing diseases.
Metagenomic methods facilitate a comprehensive, culture-independent analysis of complete microbial communities, offering vast possibilities for revolutionizing disease identification and management [4,5]. Amplicon sequencing targets conserved marker genes to characterize bacterial (16S rRNA) and fungal (ITS) communities, providing data on alpha and beta diversity [6]. The culture-independent method enables the profiling of whole communities of microorganisms. This approach permits the detection of multiple concurrent pathogens, the resolution of geographic patterns of pathogen distribution, and hidden or cryptic infections in plants that may not show symptoms or are classified as uninfected [7]. This is critical for understanding the composition and spatial distribution of bacterial and fungal pathogens. In Eritrea, so far, no comprehensive geographically based molecular systematic potato disease survey has been conducted across the diverse production zones. The absence of molecular characterization constitutes a significant gap, as such data are essential for comprehending pathogen evolution, dissemination, monitoring disease spread, understanding inoculum sources, evaluating emerging strain risks, developing better control strategies, and assisting in breeding initiatives. Characterizing pathogen populations in Eritrea will provide a significant contribution to understanding regional disease dynamics, identifying sources and hotspots, and developing coordinated regional management strategies.
2. Materials and Methods
2.1. Study Areas and Sampling
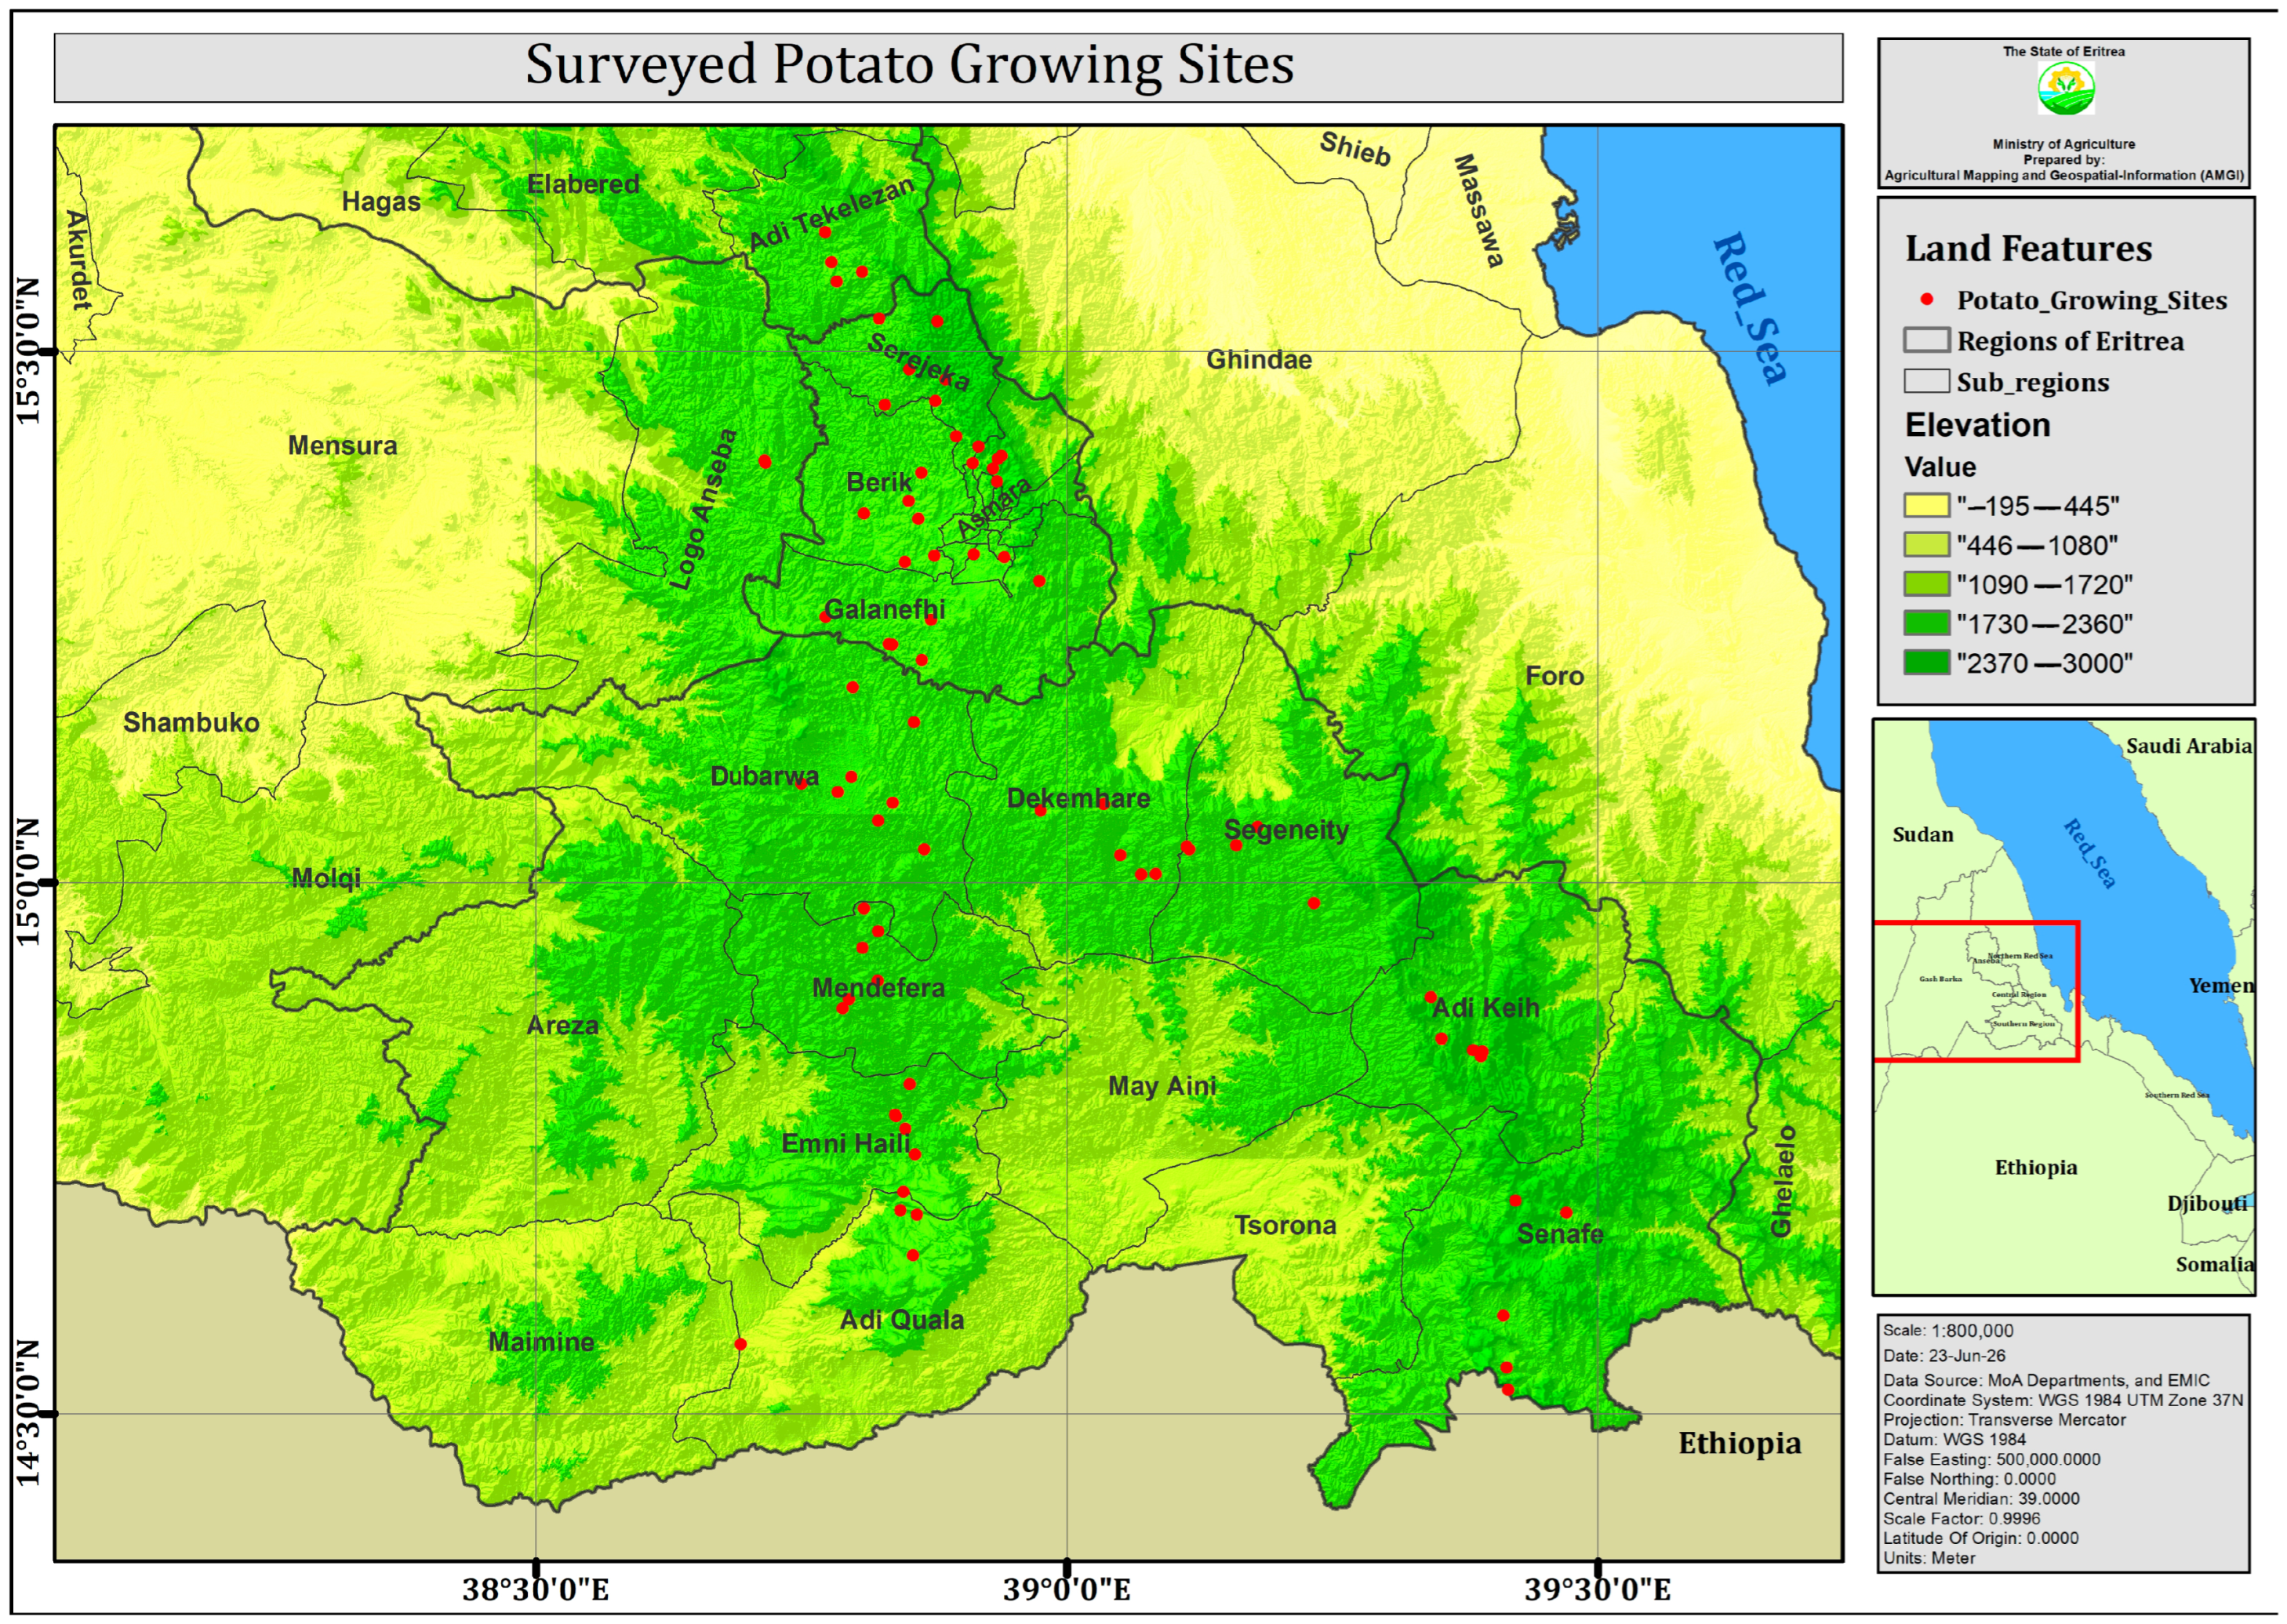
The study was conducted in 14 potato-growing sub-regions under four regions of Eritrea: Maekel (Central), Debub (Southern), Gash Barka, and Anseba ) (Figure 1). The latter two regions were represented by Logo Anseba and Adi Tekeliezan sub-regions, respectively, where their agroecology resembles the other first two major potato-growing regions. In these sub-regions, 81 farms were selected purposively to ensure comprehensive coverage of diverse agroecological zones during the growing season of July–August 2023. Moreover, the sampling sites were selected based on inclusive criteria that included major potato production areas, agroecological significance, historical disease incidence records, fields that are used for seed and ware potato production, and areas where potato production and agroecology intersected on existing maps. Additionally, clustered and pocket areas were prioritized in consultation with local sub-region extension workers.
Figure 1.
Surveyed potato growing subregions of Eritrea.
Within each selected farm, a diagonal sampling technique was used to collect leaf, stem, and tuber samples from symptomatic, diseased plants. Furthermore, representative samples were collected from asymptomatic (visually healthy-looking) plants from three potato production farms at different sub-regions, namely Dubarwa (labelled as Asy Dubarwa), Adi Keih (labelled as Asy Adi Keih), and Gala Nefhi (labelled as Asy Gala Nefhi), to investigate latent infections. Samples from the categorized diseases based on symptoms were confirmed by the National Animal and Plant Health Laboratory (NAPHL) of Eritrea based on morphology, prior to metagenomics processing. The predominant potato variety cultivated during the survey across sub-regions was Electra, a variety that is dominant in the country.
The samples collected from each farm were then pooled into one sample. Similarly, for the asymptomatic samples, one pooled sample was prepared per selected area. Each pooled sample designated for DNA extraction was immediately placed in zip-locked plastic bags. Strict disinfection protocols, including the use of ethanol and gloves, were followed for hands and tools between sampling sites to prevent cross-contamination between farms and samples.
2.2. Crude Sample Extraction
Leaf samples were ground using a pestle and mortar with liquid nitrogen at the National Animal and Plant Health Laboratory of Eritrea (NAPHL). The ground crude samples were then sealed in 2 mL Eppendorf tubes (Eppendorf SE, Hamburg, Germany), DNA/RNA Shield (Zymo Research, Irvine, CA, USA) was added to preserve nucleic acid integrity, and the samples were stored at 4 °C until further processing.
2.3. DNA Extraction
Total DNA was extracted following the protocol described by Yu and his colleagues [8] with in-house modifications at the Institute of Biotechnology Research (IBR) laboratory of JKUAT. The crude sample (300–400 µL) was washed with an equal amount of phosphate-buffered saline (PBS) by centrifuging at 13,000 rpm for 5 min to remove the DNA/RNA shield used for nucleic acid stabilization during transport. After discarding the supernatant, 0.15 g of powdered activated charcoal (PAC), 500 µL of CTAB (cetyltrimethylammonium bromide) buffer, 300 µL of SDS (sodium dodecyl sulfate) (10%), and finally 20 µL of proteinase K (20 mg/mL) were added to the 2 mL centrifuge tube. The mixture was then vortexed to homogenize and incubated in a water bath at 65 °C for 1 h with intermittent shaking. Following this, 30 µL of lysozyme was added, and samples were placed in a water bath at 37 °C for 45 min of incubation. After centrifuging at 12,000 rpm for 20 min at 4 °C, the supernatant was collected. The DNA was precipitated by adding half the amount of PEG 6000 (30% in 1 M NaCl) and left at room temperature for 1 h. Thereafter, 100 μL of 5 M potassium acetate and 1 mL of TE buffer (pH 8.0) were added, and the mixture was incubated for 15 min at 4 °C. After adding an equivalent volume of phenol-chloroform-isoamyl alcohol (25:24:1), the mixture was centrifuged at 10,000 rpm for 10 min at 4 °C after standing at room temperature for 10 min. The aqueous layer was transferred into a new tube, treated with 0.7 volume of isopropanol, and left for 1 h at room temperature. DNA was precipitated by centrifugation at 10,000 rpm for 10 min at 4 °C. The final pellet was centrifuged at 10,000 rpm for 10 min at 4 °C and dried at room temperature after being successively cleaned with 1 mL of 75% ethanol and 1 mL of anhydrous alcohol. The dried pellet was eluted in 60 µL of nuclease-free water and stored at −20 °C until further analysis. The collected DNA was subsequently pooled into 17 samples that represented the country’s sampled potato-growing sub-regions (based on 14 symptomatic + 3 asymptomatic samples during collection) [9].
2.4. Library Preparation and Sequencing
Using the Nextera XT Index V2 Kit (Illumina Inc., San Diego, CA, USA), the amplicon library was built in accordance with the procedures for 16S and ITS Metagenomic Sequencing Library preparation. The internal transcribed spacer (ITS1 and ITS2) region was targeted for fungal community profiling, whereas the V3–V4 hypervariable region of the 16S rRNA gene was amplified for bacterial community profiling. The adapter and PCR primers utilized are described in Table 1. Paired-end sequencing (2 × 301 bp) was performed on an Illumina MiSeq (Illumina Inc., San Diego, CA, USA). A total of 34 libraries were sequenced, comprising 17 for 16S rRNA and 17 for ITS regions.
Table 1.
Primers and adaptors used in sequencing on the Illumina MiSeq platform.
2.5. Data Preprocessing and ASV Generation
Following sequencing, adapter and primer sequences were removed from the raw data using Cutadapt (v3.2) [10]. Additionally, Cutadapt (v3.2) was used to trim forward and reverse reads to 250 bp and 200 bp. Amplicon Sequence Variants (ASVs) were generated through error-correction, merging, and denoising processes implemented in DADA2 (v1.18.0) [11]. Sequences with an expected error rate of 2 or more were excluded. Erroneous reads were denoised based on an established error model, and paired-end reads were merged by overlapping. Chimeric sequences were removed using the consensus method using the removeBimeraDenovo function in DADA2, and ASVs shorter than 350 bp were filtered out using R (v4.0.3). QIIME 2 (v2024.2) [12] was used for normalization in the microbial community comparison analysis. To ensure comparability, subsampling was carried out based on the sample with the lowest read count across all samples. The resulting ASVs were utilized for downstream analysis.
2.6. Taxonomy Analysis and Community Diversity
The organism with the highest resemblance in the relevant reference database (NCBI BLASTn) was matched to each ASV. The taxonomic and community diversity were then analyzed using QIIME 2 (v2024.2) [12]. To understand species complexity within individual samples, alpha diversity metrics such as the Shannon index, Gini–Simpson index, and PD Whole tree were computed. Bray–Curtis, Weighted UniFrac, and Unweighted UniFrac distances were used to calculate beta diversity, or the variation among samples within a group. Genetic relationships among samples were visualized based on Principal Coordinates Analysis (PCoA) and Unweighted Pair Group Method with Arithmetic Mean (UPGMA) tree.
3. Results
3.1. Sequencing and ASV Output
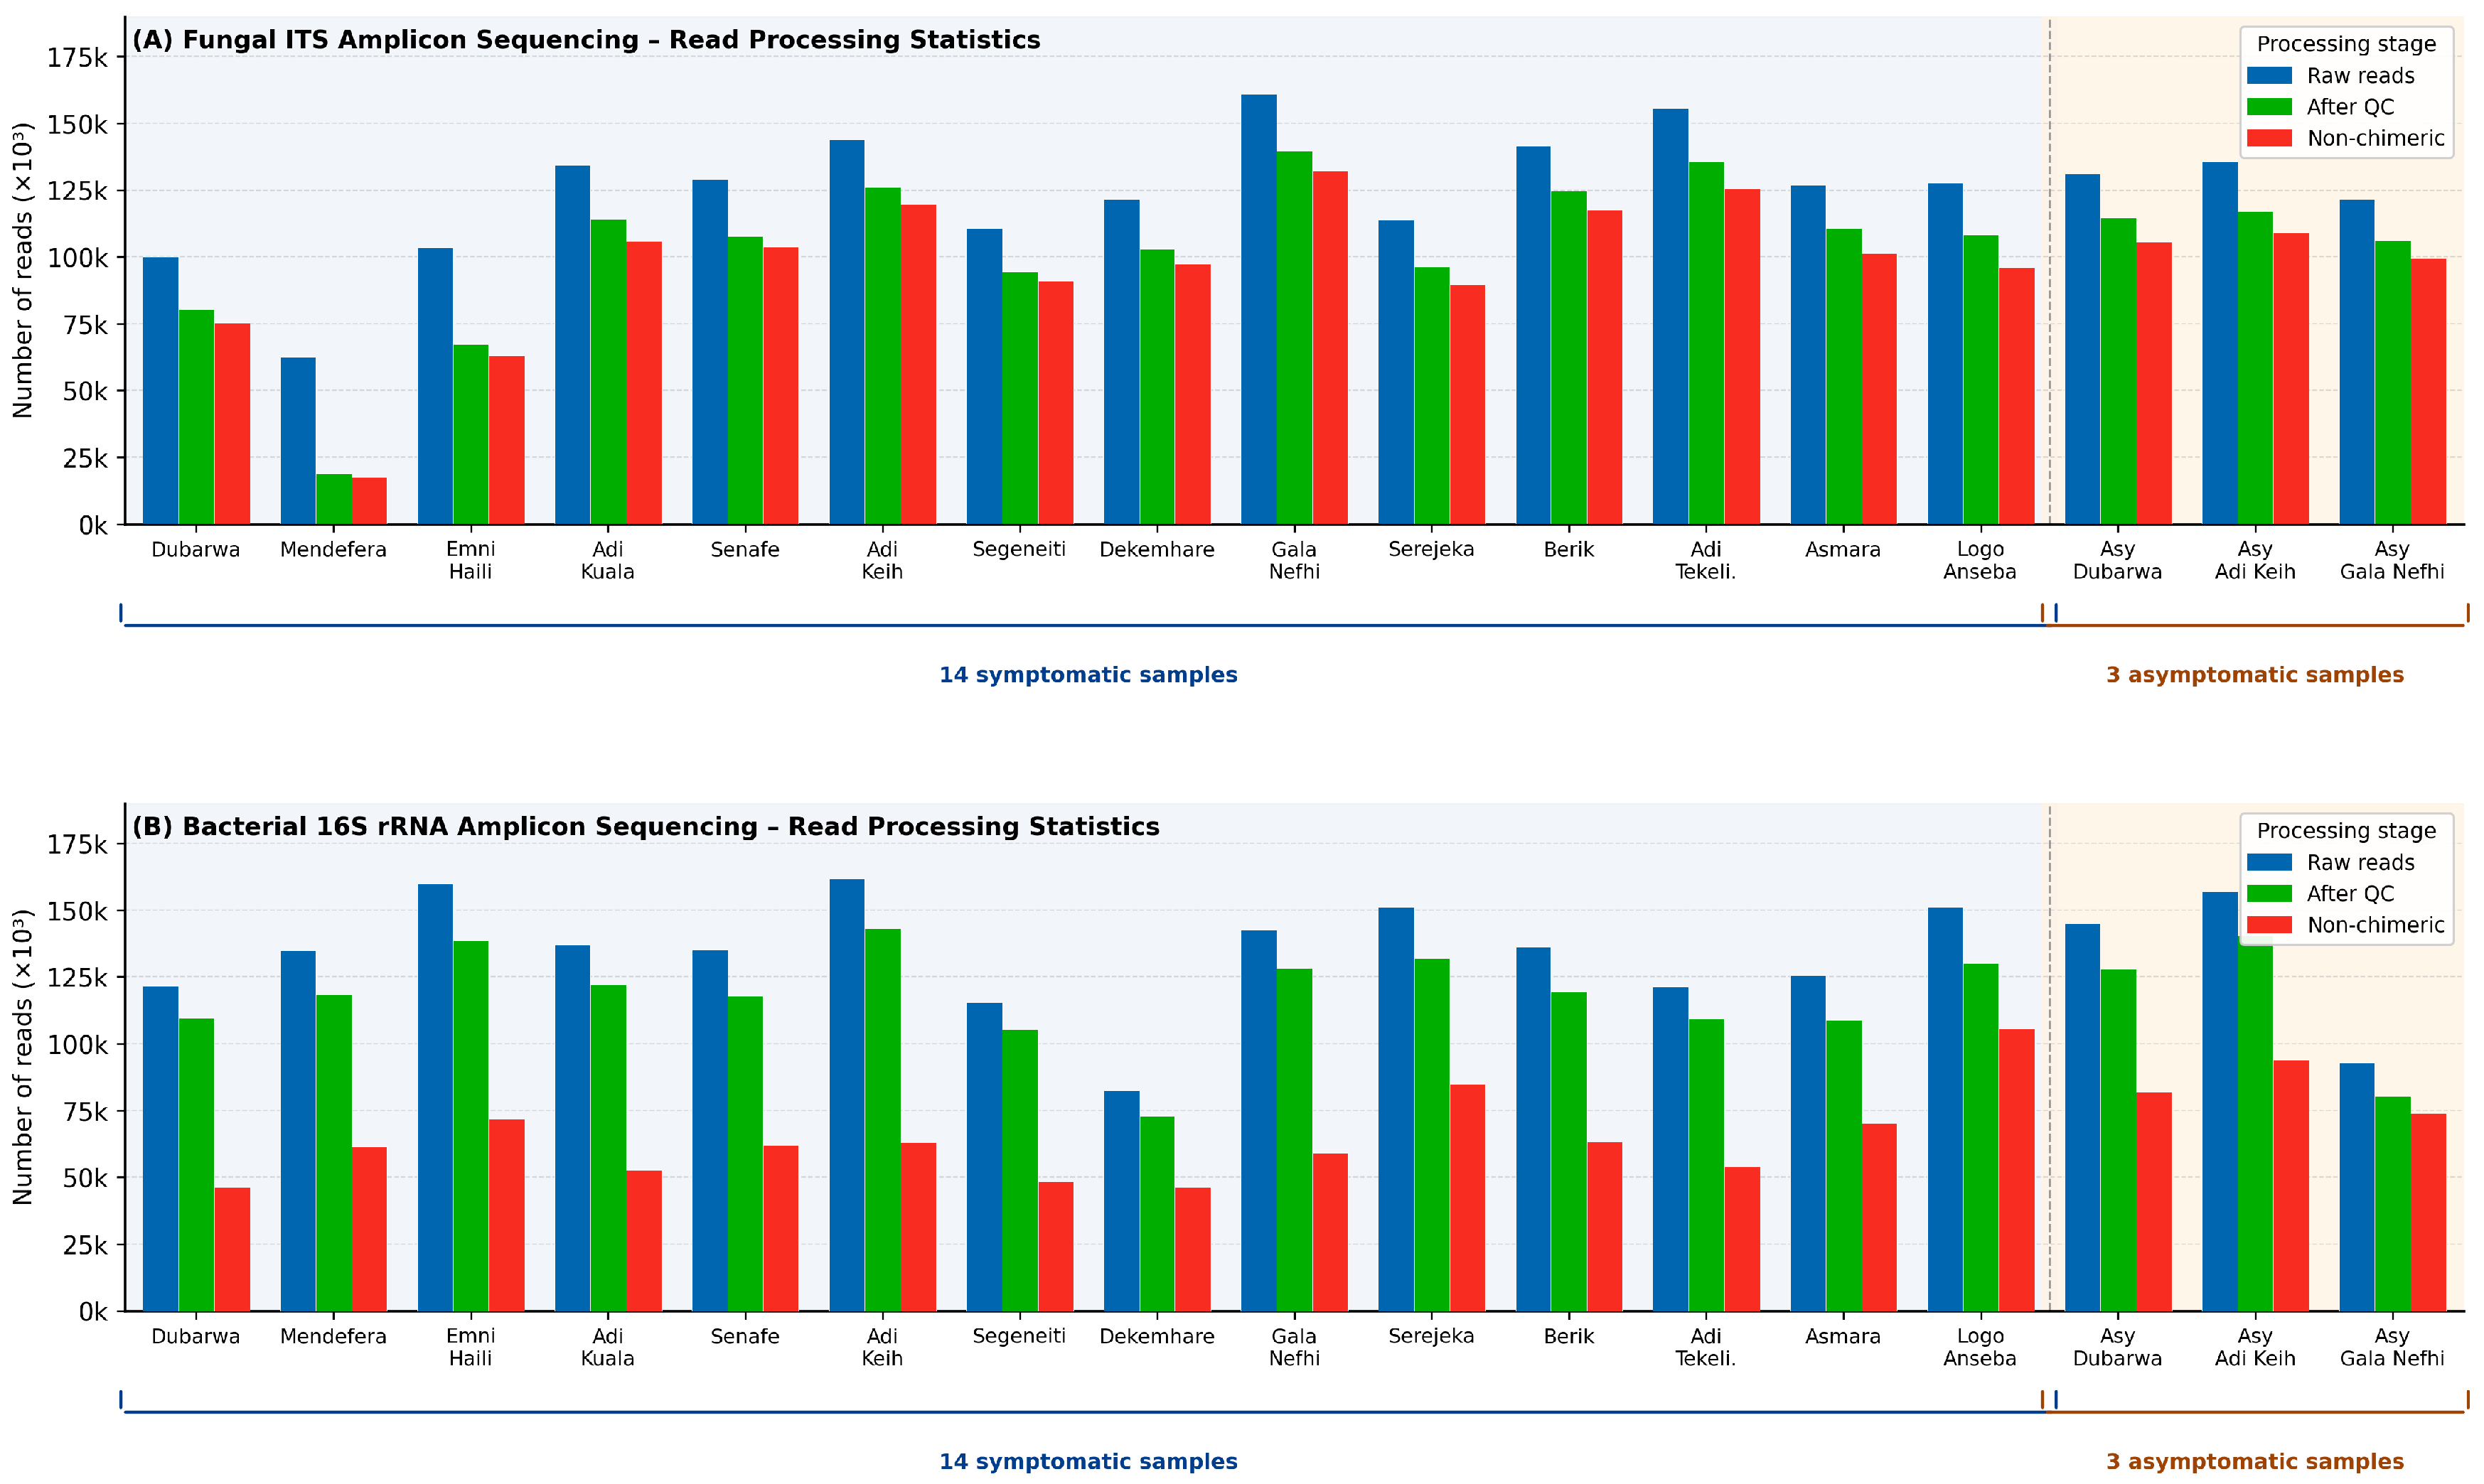
Quality filtering retained 110,000–155,000 bacterial reads per sample; after denoising and chimera removal, 50,000–110,000 non-chimeric reads per sub-region were retained for ASV generation. Fungal sequences showed similar retention (40,000–90,000 final reads), providing sufficient depth for diversity analysis. Asymptomatic samples (Asy Dubarwa, Asy Adi Keih, Asy Gala Nefhi) showed lower read counts, reflecting lower microbial compositions. The DADA2 pipeline successfully denoised sequences and removed chimeras, generating high-quality ASV tables. Bacterial communities yielded between 119 and 536 unique ASVs across locations (median = 391 ASVs), while fungal communities contained 84 to 266 ASVs (median = 168 ASVs) (Figure 2).
Figure 2.
Quality control read processing statistics for fungal ITS (A) and bacterial 16S rRNA (B) amplicon sequencing. Bars represent read counts at three preprocessing stages: raw reads, after quality control, and final non-chimeric reads across 17 samples.
3.2. Geographic Distribution of Bacterial Pathogens
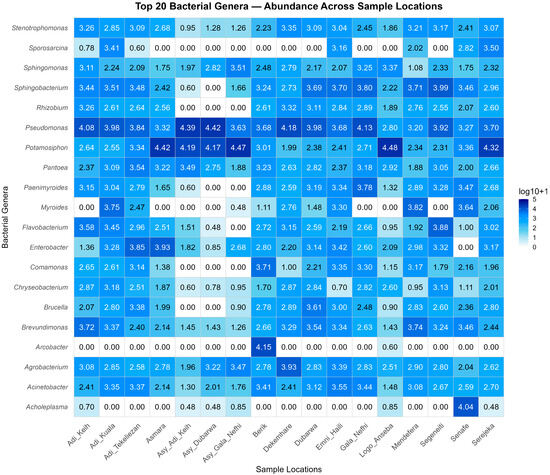
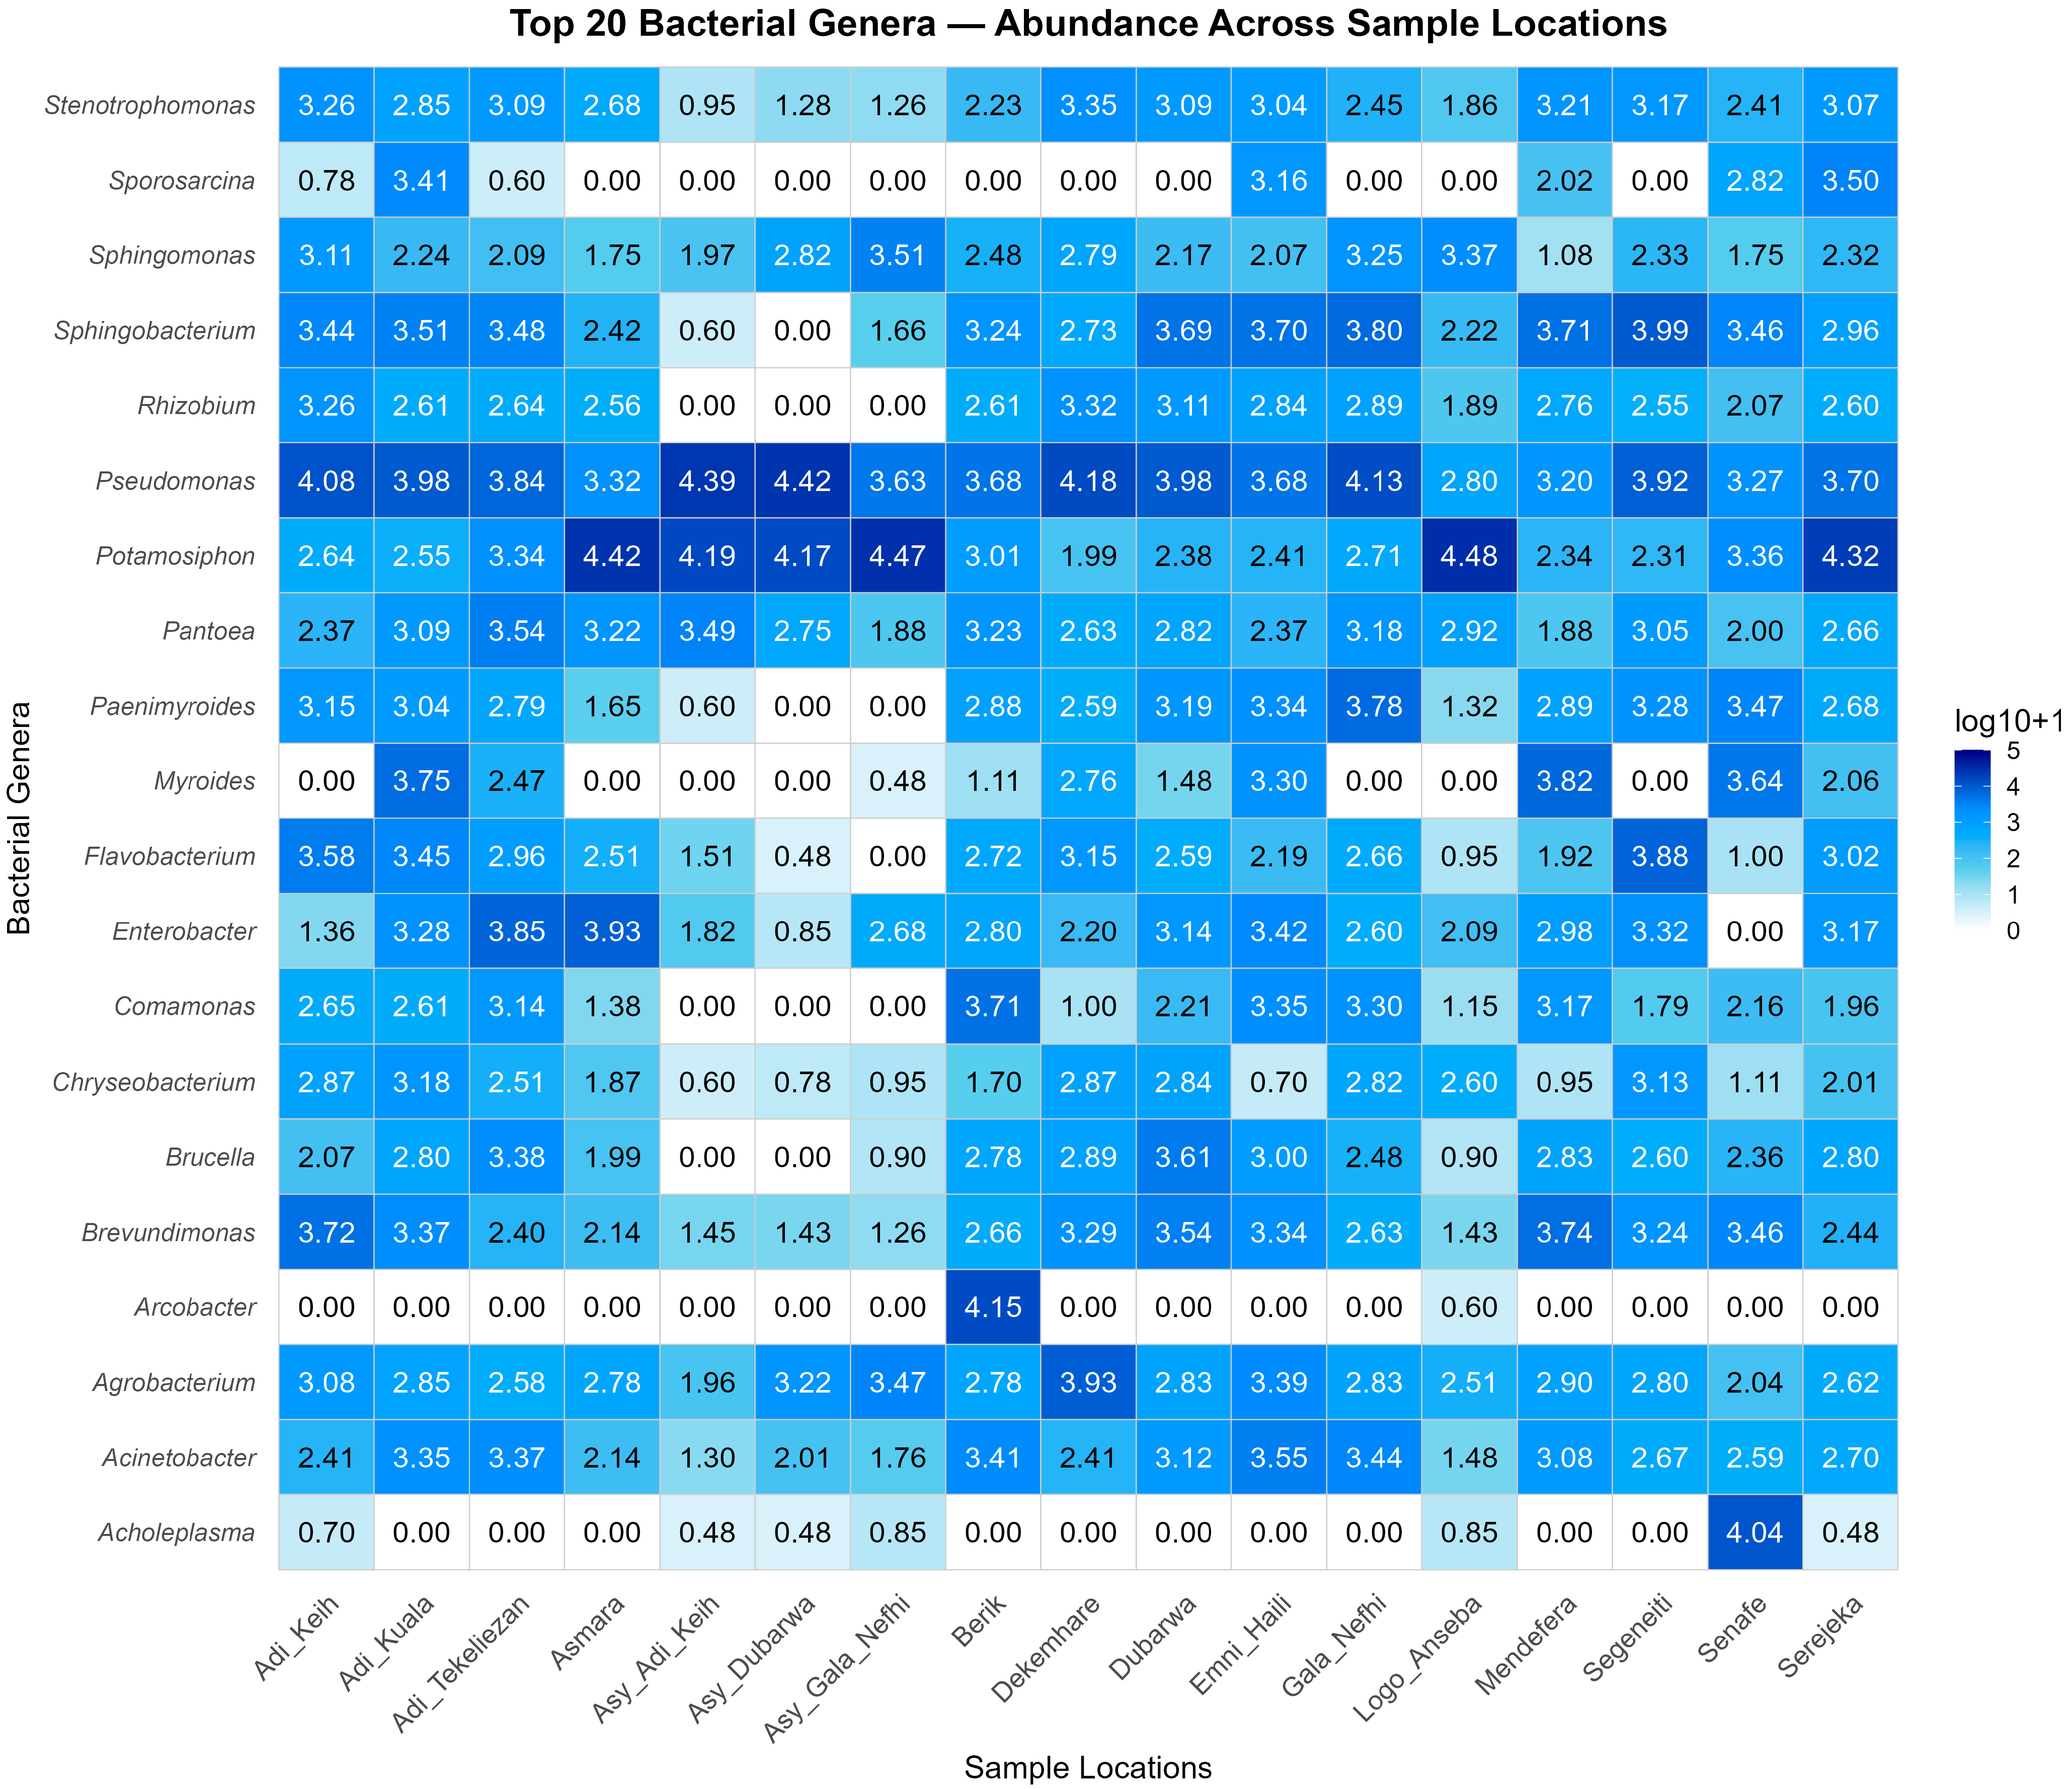
Taxonomic profiling of the samples revealed substantial geographic variation in bacterial communities across the 14 sub-regions (Figure 3). Pseudomonas species were detected in all locations, exhibiting varying relative abundances in the sampling sites. The highest Pseudomonas abundances were observed in Dekemhare (32.5%), Gala Nefhi (29.3%), Adi Keih (25.8%), Dubarwa (20.8%), and Adi Kuala (20.8%), while lower abundances were recorded in Logo Anseba (1.4%), Mendefera (3.4%), Senafe (4.0%), and Asmara (4.6%).
Figure 3.
Geographic distribution of major bacterial genera across fourteen potato-producing sub-regions in Eritrea.
Pectobacterium, the causal agent of economically important bacterial soft rot and blackleg diseases, exhibited different geographic distribution patterns. Higher Pectobacterium abundances were detected in Berik (3.3%), followed by Adi Tekeliezan (0.9%). The predominant species included P. atrosepticum, P. carotovorum, P. quasiaquaticum, P. aroidearum, P. polaris, and P. betavasculorum. Moreover, P. atrosepticum was observed to be more common in cooler highland locations in Adi Keih, whereas P. carotovorum was more abundant in Berik. Ralstonia solanacearum, the bacterial wilt pathogen, was detected at low abundance (1.33%) in the sub-region at the cooler highland (Senafe) at one farm located at the border. Other bacterial genera detected at lower abundances included Erwinia and Xanthomonas, all of which contain species capable of causing disease in solanaceous crops.
3.3. Geographic Distribution of Fungal Pathogens
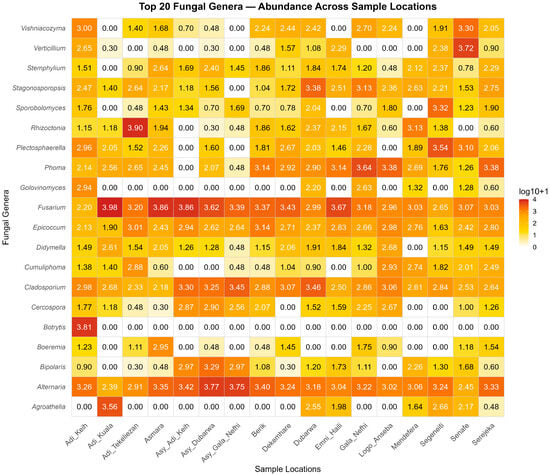
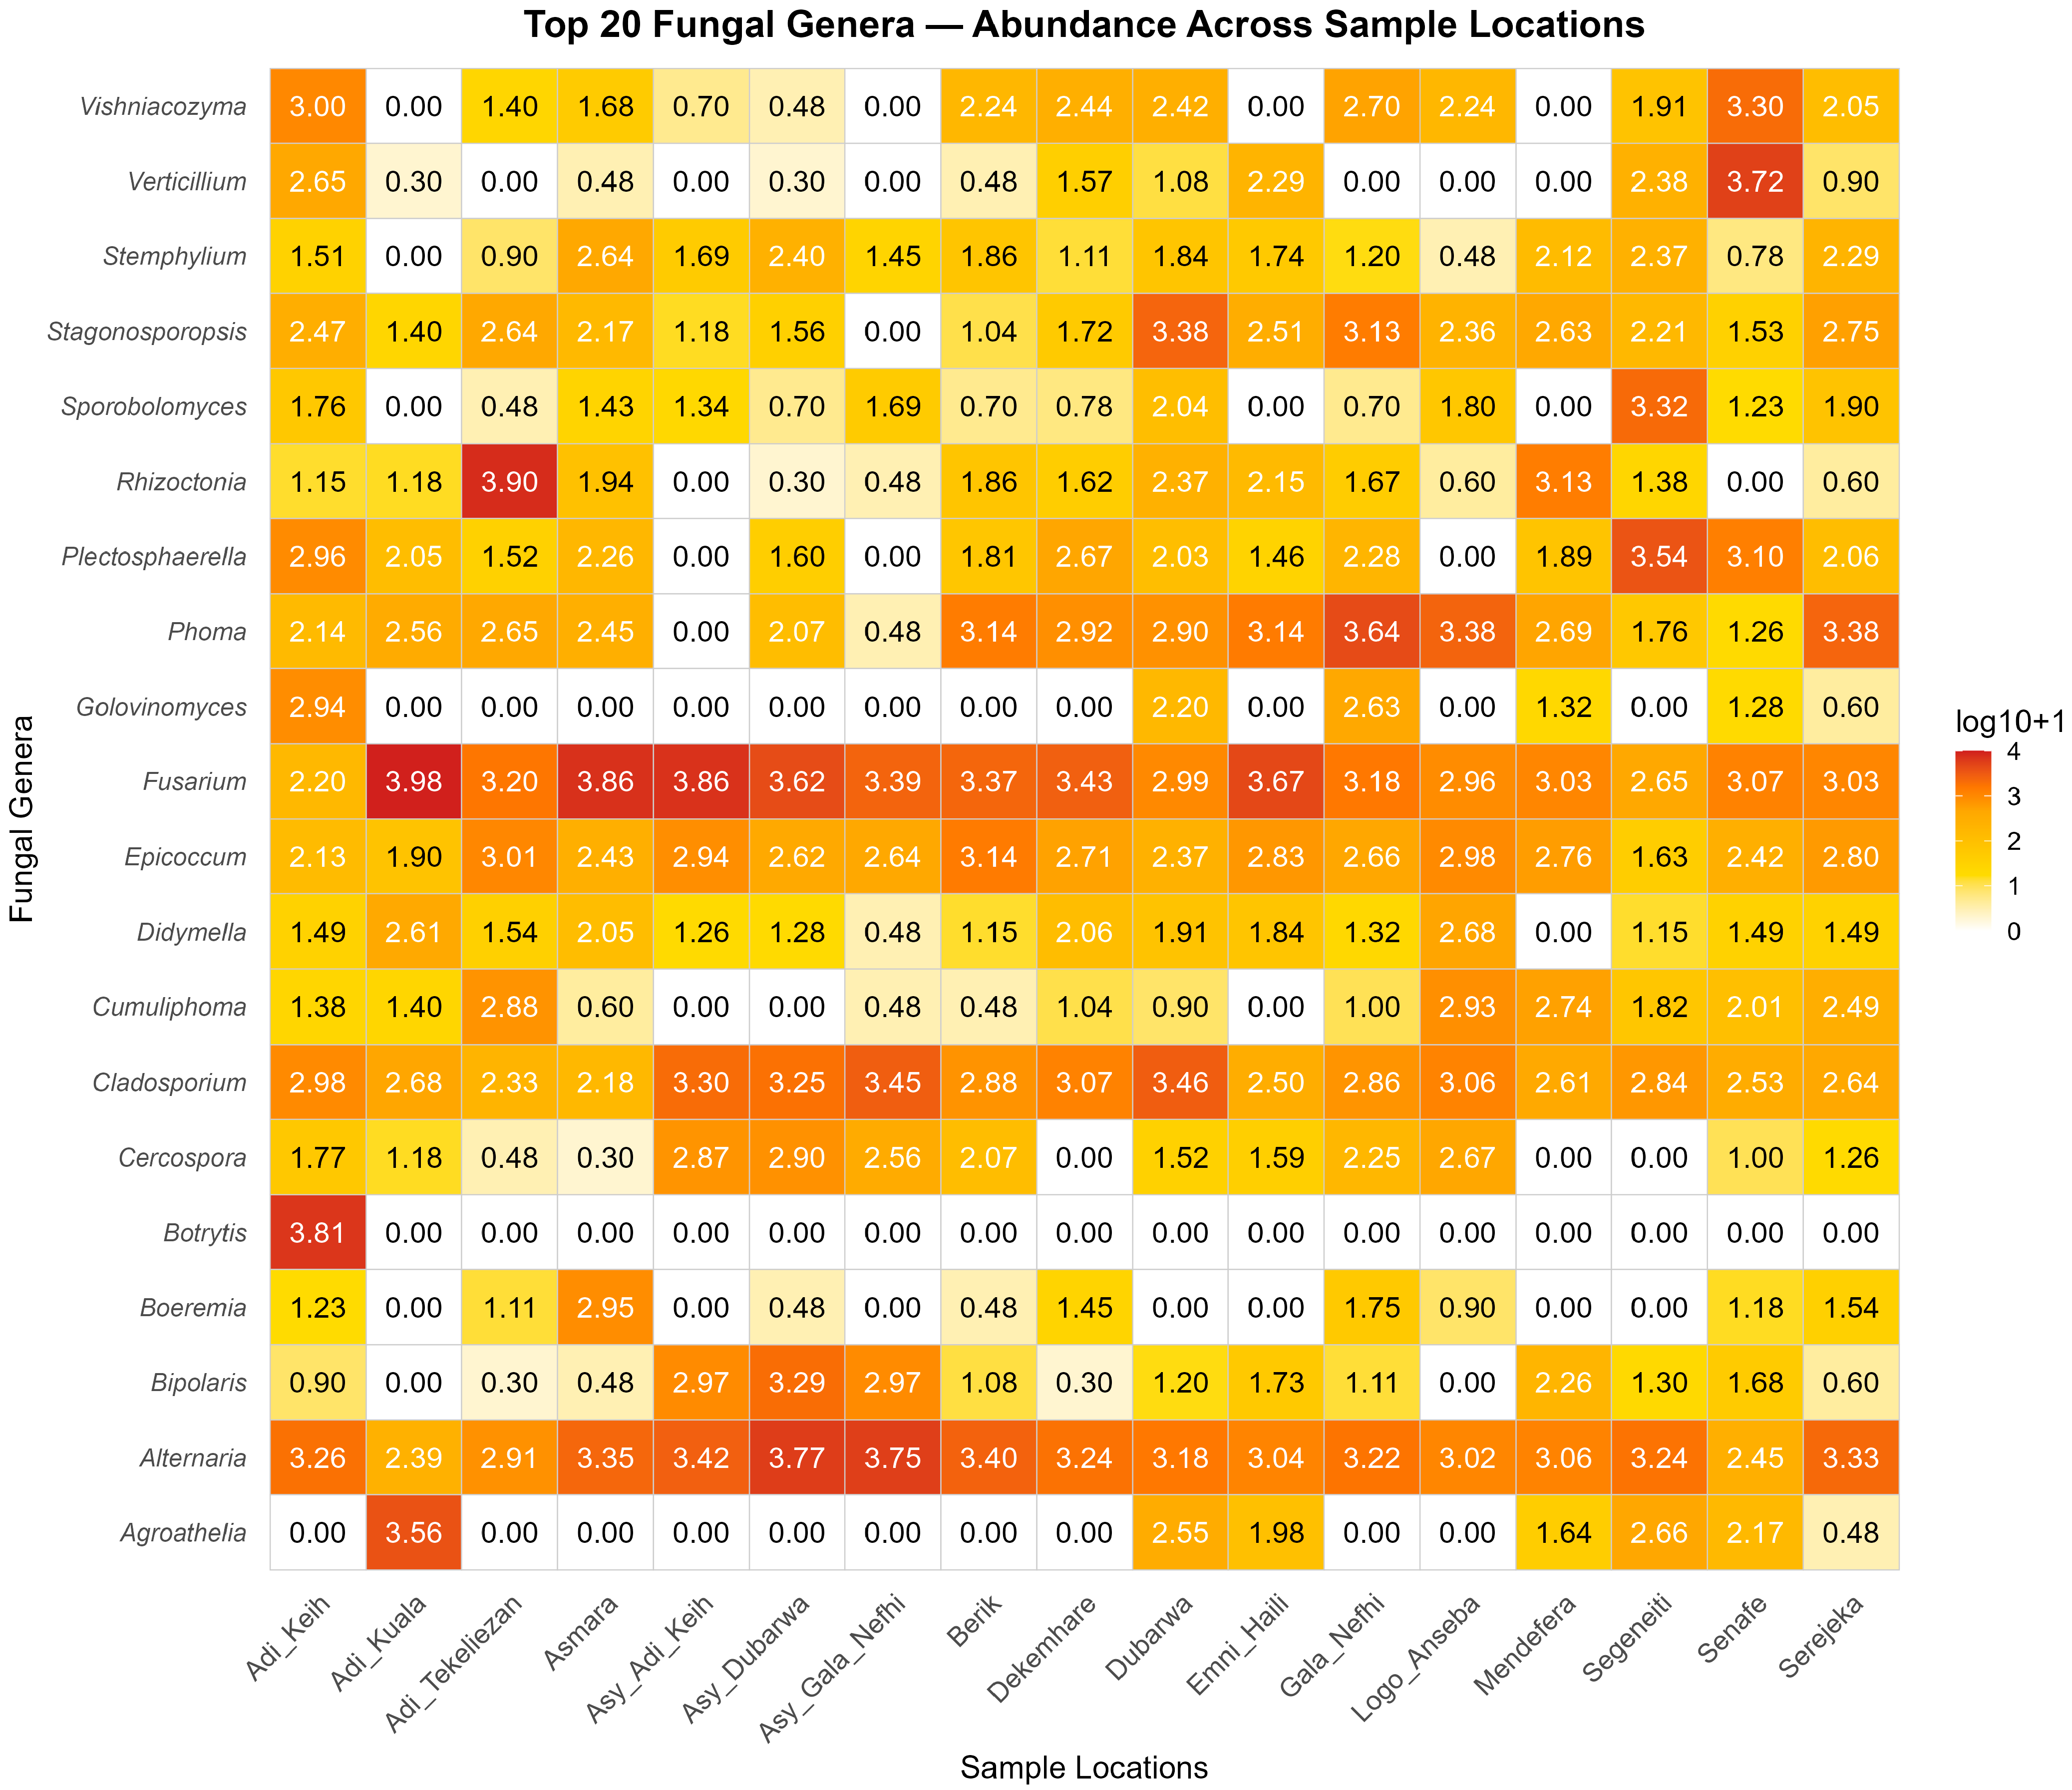
Fungal communities also showed pronounced geographic structuring across the sub-regions (Figure 4). Alternaria species, the causal agents of early blight disease, were among the most abundant fungal pathogens detected, with particularly high relative abundances in Berik (14.3%), Asmara (12.7%), and Serejeka (12.2%). Lower Alternaria abundances were observed in Senafe (1.6%), Adi Kuala (1.4%), and Adi Tekeliezan (4.5%). The multiple Alternaria spp. ASVs identified include A. solani, A. alternata, and A. tenuissima. Fusarium species, known to cause dry rot, wilt, and tuber decay in potatoes, showed presence across all locations. Fusarium community composition varied geographically, with the highest diversity observed in Adi Kuala (53.8%), Asmara (40.8%), and Emni Haili (26.2%). Species-level analysis revealed sequences matching F. solani and F. oxysporum as predominant in most locations.
Figure 4.
Geographic distribution of major fungal genera abundance across fourteen potato-producing sub-regions in Eritrea.
Botrytis cinerea, the grey mould pathogen, was detected at higher abundance (36.9%) in Adi Keih, a location characterized by higher humidity and cooler temperatures. Rhizoctonia, causal agents of stem canker and black scurf, and Verticillium, causing potato early dying, were each detected across multiple sub-regions at abundances ranging from 2.5–44.4%. Meaningful abundance (≥2%) of Rhizoctonia occurs at only two sites: Mendefera (7.6%) and Adi Tekeliezan (44.4%), and Verticillium was higher at Senafe (29.9%).
3.4. Alpha Diversity Patterns Across Geographic Locations
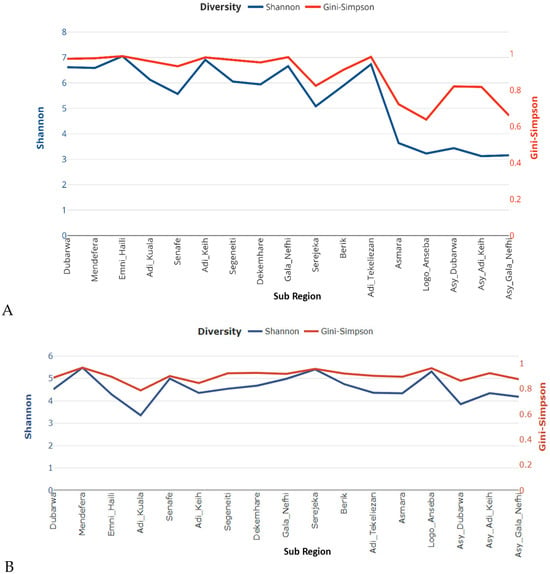
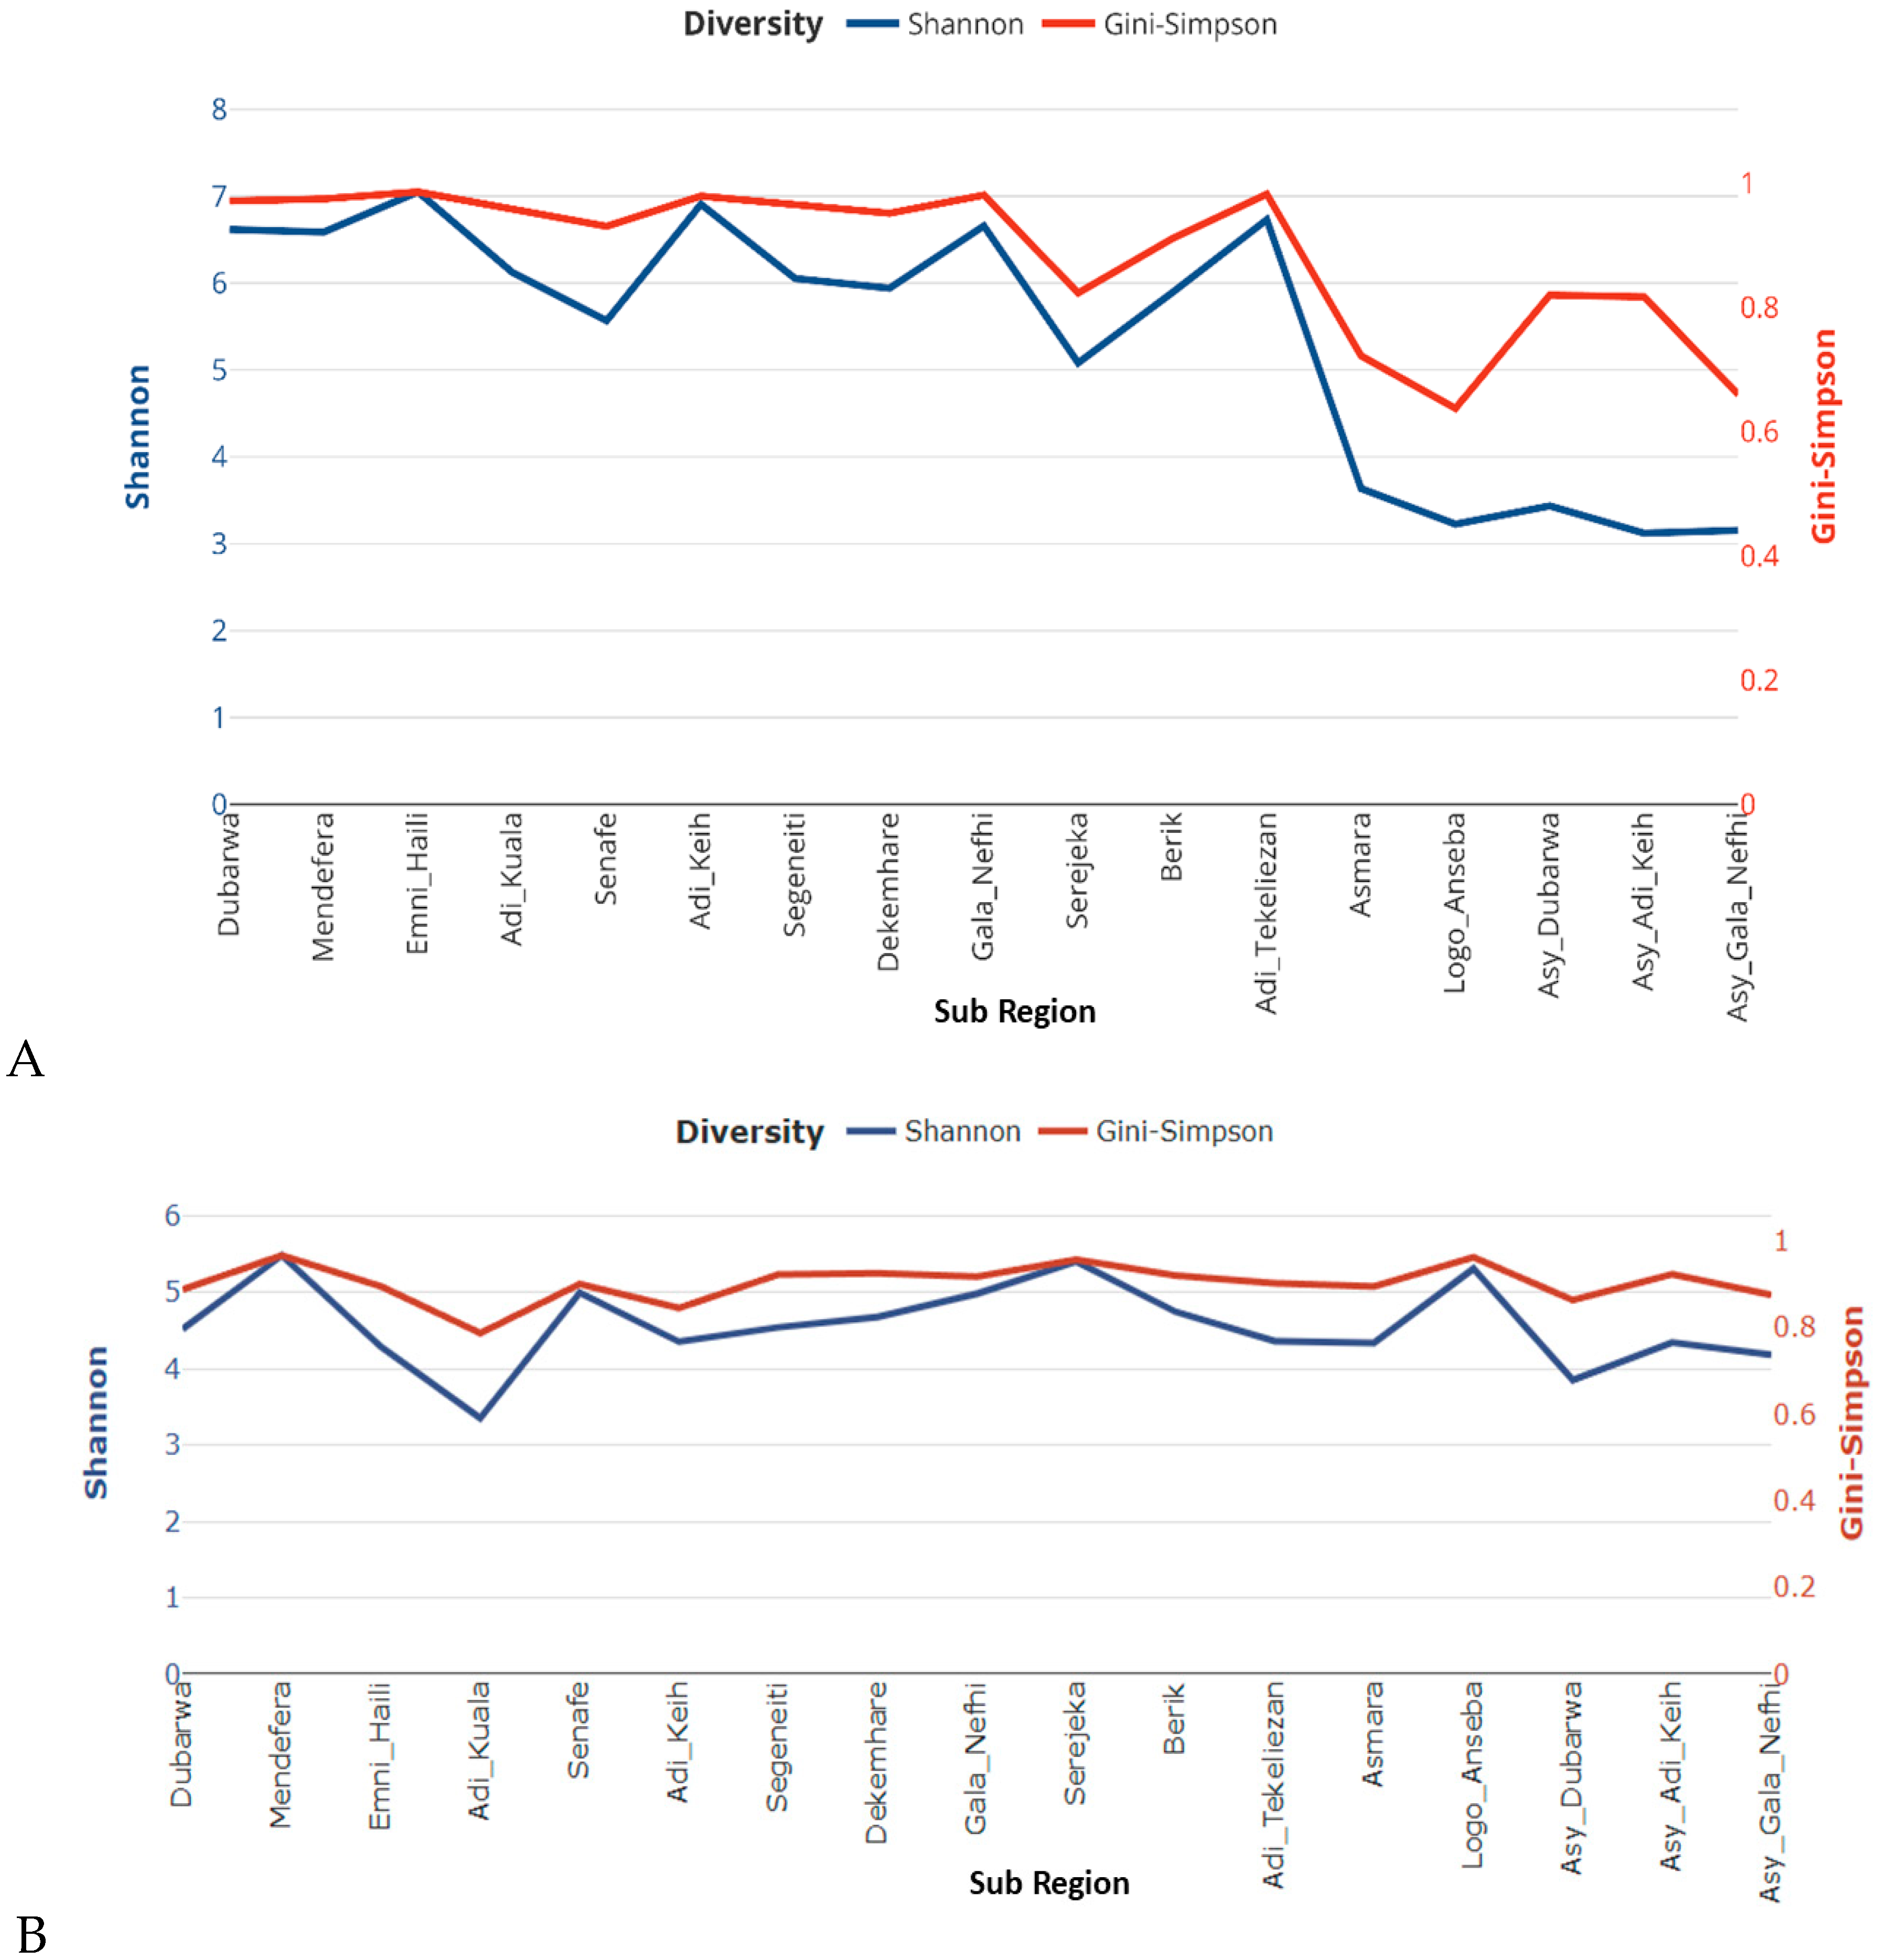
Alpha diversity analysis revealed substantial geographic variation in pathogen community complexity (Figure 5, Table 2). Bacterial Shannon diversity indices ranged from 3.12 in the asymptomatic sample from Gala Nefhi to 7.05 in the symptomatic sample from Emni Haili, with a mean of 5.67 across the 14 diseased sample locations. High bacterial diversity was found in Emni Haili (7.05), Adi Keih (6.91), Dubarwa (6.62), and Gala Nefhi (6.66). Conversely, lower diversity was observed in Senafe (5.57), Serejeka (5.08), and Dekemhare (5.94). The three asymptomatic samples showed lower diversity (mean Shannon index = 3.36).
Figure 5.
Alpha diversity patterns of bacterial (A) and fungal (B) communities across all seventeen samples.
Table 2.
Complete diversity index values for all samples.
Fungal diversity showed less geographic variation among diseased samples, ranging from 3.36 (Adi Kuala) to 5.49 (Mendefera), with a mean of 4.70. The highest fungal diversity was observed in Mendefera (5.49), Logo Anseba (5.31), and Serejeka (5.41), while the lowest was in Adi Kuala (3.36) and the asymptomatic sample from Dubarwa (3.85). Phylogenetic diversity (Table 2) patterns generally paralleled Shannon diversity values, ranging from 12 to 40 for bacteria and 22 to 40 for fungi.
3.5. Geographic Clustering of Pathogen Communities
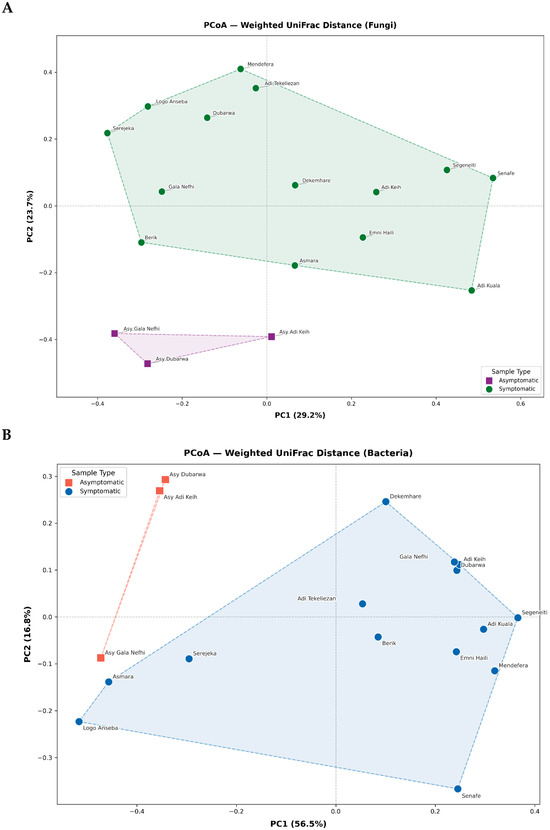
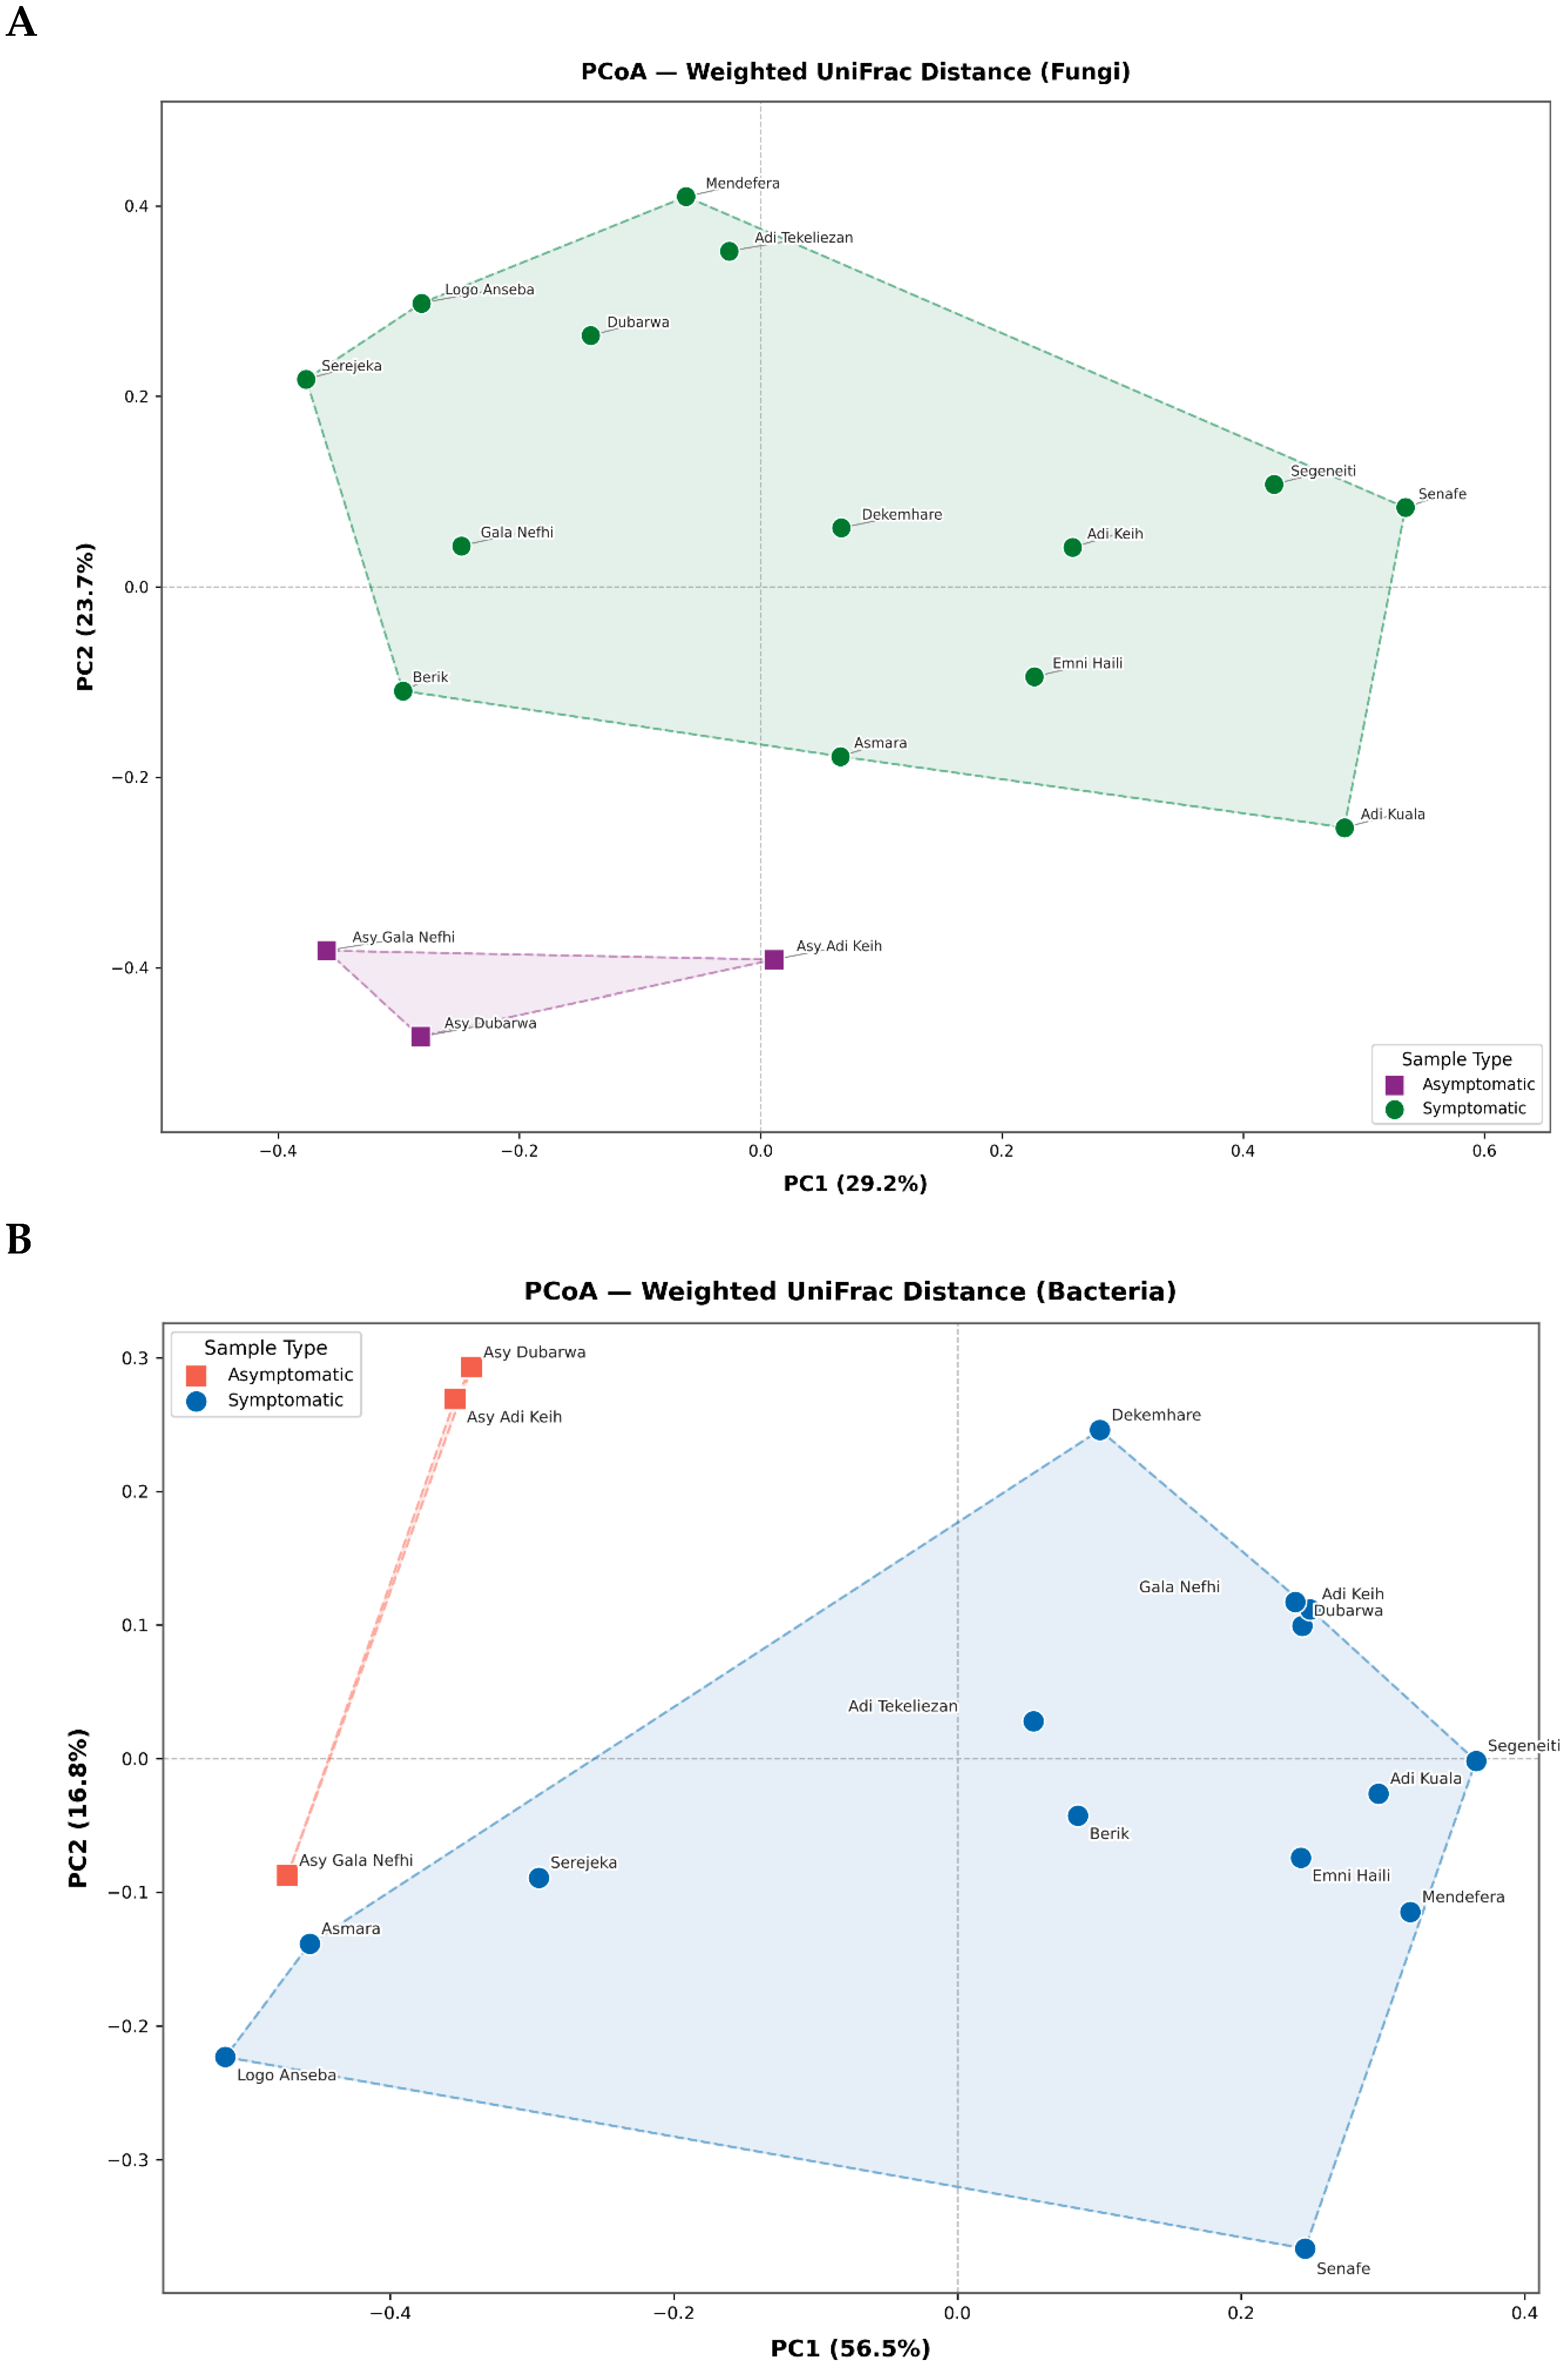
Beta diversity analysis revealed distinct geographic structuring of pathogen communities (Figure 6). Principal Coordinates Analysis (PCoA) based on Bray–Curtis dissimilarity explained 27.91% (PC1) and 12.28% (PC2) of the variance in bacterial communities, and 16.97% (PC1) and 13.61% (PC2) in fungal communities. For bacterial communities, Senafe and Serejeka showed clear separation from other locations along PC1, suggesting distinctive microbial community profiles in these sub-regions. Berik, Gala Nefhi, and Dekemhare formed a loose cluster, possibly reflecting similar environmental conditions or disease management practices. The three asymptomatic samples clustered together and separately from most diseased samples. Weighted UniFrac PCoA, which incorporates phylogenetic relationships, showed even stronger patterns, with PC1 explaining 56.5% of bacterial community variance (Figure 6B). Fungal weighted UniFrac PCoA explained 29.25% (PC1) and 23.73% (PC2) of variance (Figure 6A), displaying similar geographic clustering patterns.
Figure 6.
PCoA ordination plots of (A) fungal communities and (B) bacterial communities based on weighted UniFrac.
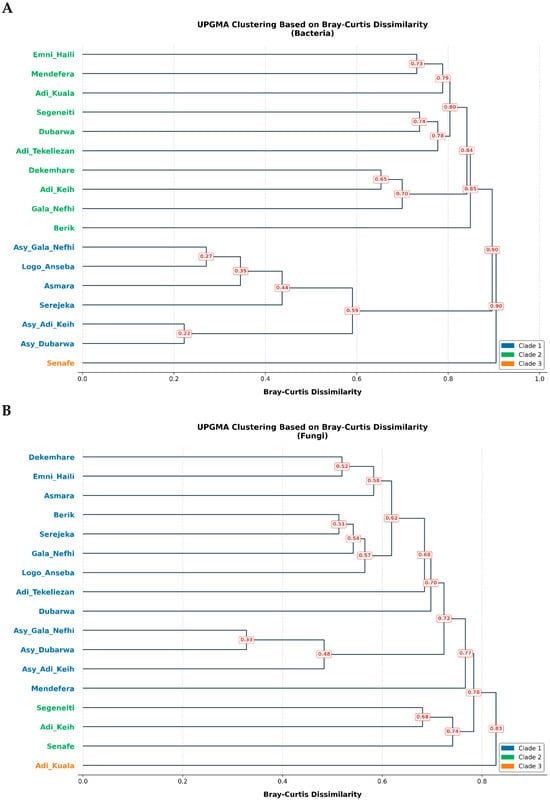
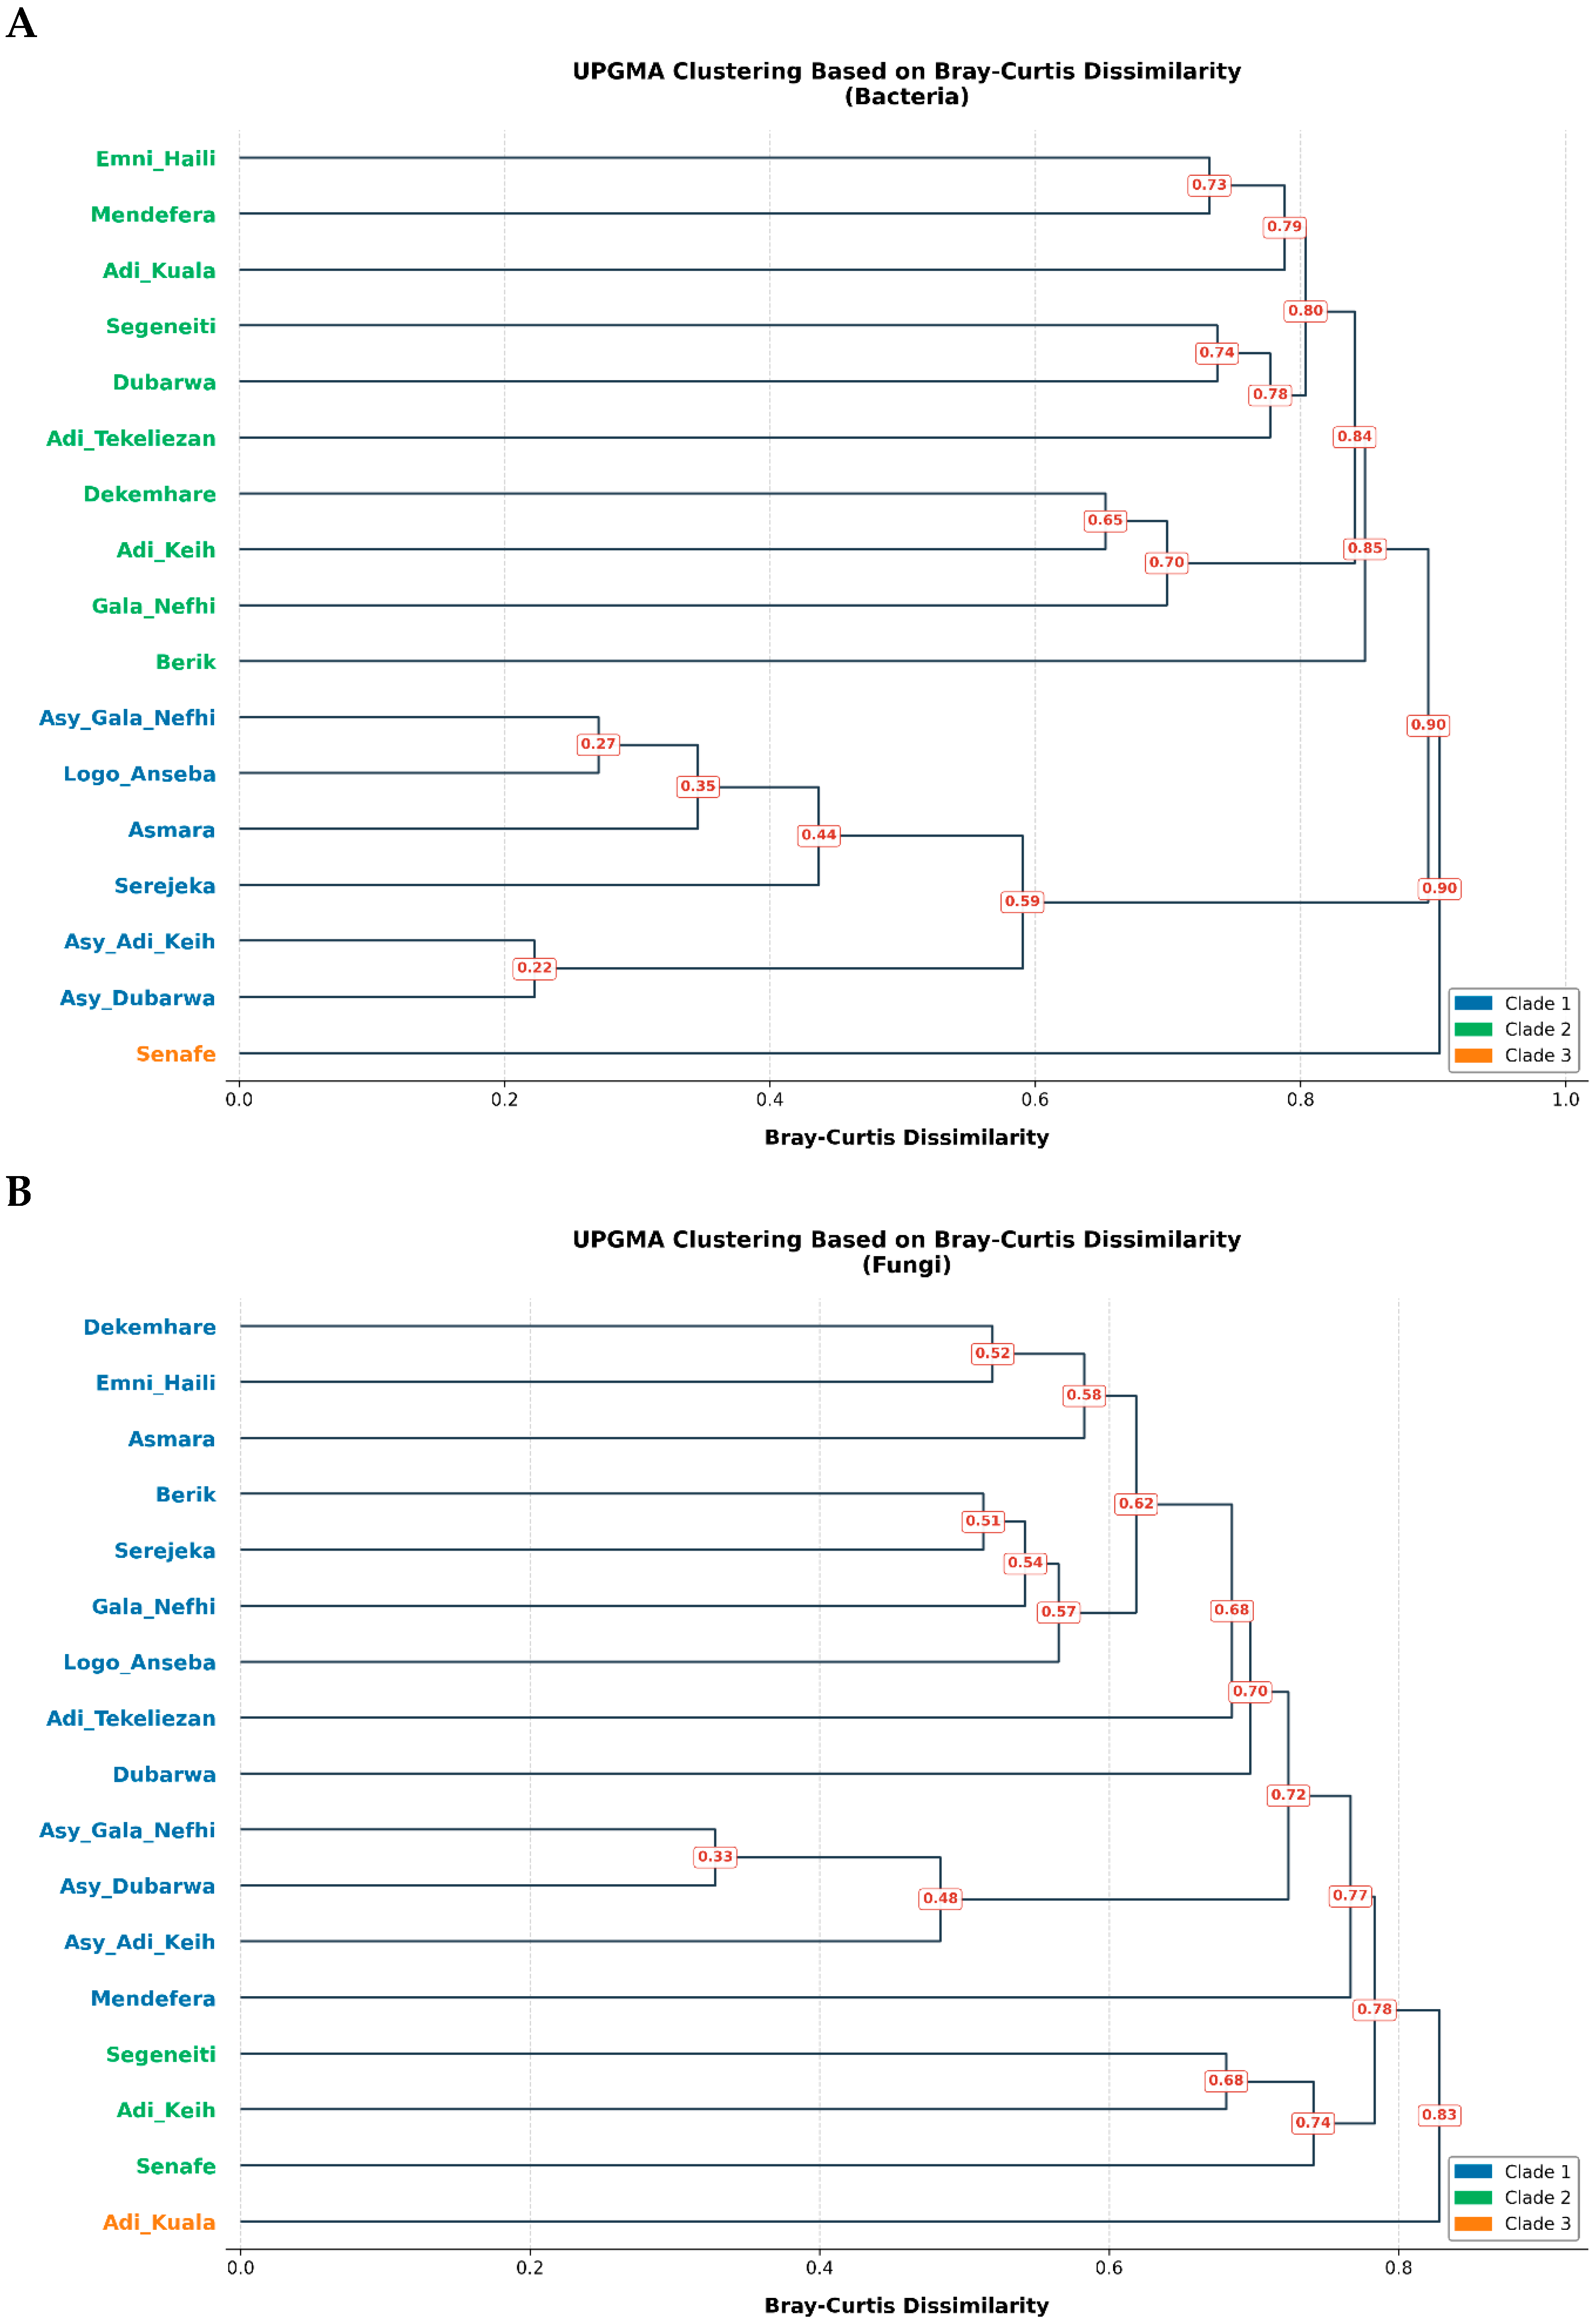
UPGMA clustering based on Bray–Curtis dissimilarity produced dendrograms that reflected geographic proximity. For bacteria, three main clades were identified, where Senafe was the most divergent sub-region, joining all others last, while for fungi, Adi Kuala was the most divergent sub-region (Figure 7). Distance matrix analysis revealed Bray–Curtis dissimilarities ranging from 0.22 to 0.90 among sub-regions for bacteria and 0.33 to 0.83 for fungi. For bacteria, the lowest dissimilarities were observed between geographically similar sub-regions, Adi Keih–Dekemhare (0.65) and Adi Keih–Gala Nefhi (0.70), in the Debub and Mekel regions, respectively; for fungi, Serejeka-Berik (0.51) and Emni Haili-Dekemhare (0.52) showed closer dissimilarity. The highest dissimilarities occurred between geographically distant sub-regions, including Mendefera–Logo Anseba (0.97) and Dubarwa–Logo Anseba (0.96) for bacteria, and Adi Kuala–Adi Keih (0.90) and Adi Kuala–Asy Gala Nefhi (0.88) for fungi (Table S1).
Figure 7.
Weighted UniFrac UPGMA clustering based on Bray–Curtis Dissimilarity. (A) Sub-region clustering based on bacterial communities; (B) sub-region clustering based on fungal communities.
3.6. Pathogen Detection in Asymptomatic Plants
The samples collected from asymptomatic potato plants showed a consistent presence of pathogen-associated taxa in all three asymptomatic samples, albeit at low abundances. Bacterial pathogens associated taxa detected in these asymptomatic samples included Pseudomonas, and similarly, fungal pathogens associated with the taxa identified included Alternaria, Fusarium, and Colletotrichum.
4. Discussion
The characterization of microbial communities associated with diseased potato (Solanum tuberosum L.) across the potato growing sub-regions of Eritrea provides a comprehensive and ecologically grounded picture of the disease complex affecting potato production in the country. The high-throughput amplicon sequencing approach identified a unique microbial landscape, revealing significant geographic structuring and highlighting the need for advanced surveillance in disease monitoring and seed systems. In East African potato-growing highlands, surveys conducted in Kenya, Uganda, and Ethiopia have identified Ralstonia solanacearum, Fusarium solani, Alternaria solani, and Pectobacterium spp. as the dominant disease agents [13,14,15], consistent with the community composition detected in the present study. The occurrence of these shared taxa across the broader East African highland potato belt underscores the regional coherence of potato disease pressure and the relevance of coordinated surveillance strategies. In the Andean highlands, the centre of origin of potato, metagenomic surveys of potato rhizosphere microbiomes have similarly documented site-dependent variation in pathogen community composition driven by altitude and vegetation stage [16], mirroring the geographic structuring observed here across Eritrea’s altitudinal gradient. In Canadian potato systems, amplicon-based surveys have identified Verticillium dahliae and Rhizoctonia solani as dominant soilborne threats [17], a pattern that parallels their prominence in the cooler, higher-altitude sub-regions of Eritrea detected in this study.
The widespread occurrence of Alternaria species, particularly A. solani and A. alternata, indicates that early blight is a major and persistent threat to Eritrean potato production. The low Bray–Curtis dissimilarity for Alternaria across sub-regions suggests an established inoculum reservoir, implying that preventive, weather-based fungicide applications are more effective than reactive treatments [18,19]. The geographic structuring of pathogen communities is in agreement with a study conducted by Davies and Pedersen [20], where the pathogen communities are influenced by host range and geographical clustering. Where host relatedness is the best overall predictor of whether two host species share the same pathogens, and geographical overlap among neighbouring hosts is more important in determining host range [20]. The dominance of Pseudomonas and Pectobacterium as major bacterial pathogens, and Alternaria and Fusarium species as major fungal pathogens, is in line with their global significance in potato production [21,22]. In a similar recent study of metagenomic characterization of bacterial abundance related to potato by Kiige and his team [23] found that Enterobacteriaceae and Pseudomonadaceae were the most dominant bacterial families present in the rhizosphere. The identification of the spread of Pectobacterium species, P. atrosepticum in cooler highlands and P. carotovorum in warmer highland sub-regions aligns with the characteristics of the bacteria, where moist, cool (below 70 °F or 21 °C) conditions enhance blackleg caused by P. atrosepticum, while warmer conditions (70 °F to 80 °F or 21–27 °C) are optimal for soft rot caused by P. carotovorum [24]. This indicates that local climate is an important factor contributing towards the distribution of particular pathotypes [25] and thus has potential implications for the development of region-specific management strategies.
The detection of Verticillium dahliae in varying abundance is epidemiologically significant. The pathogen infects roots, blocks vascular tissues, and leads to premature vine yellowing and wilting, thereby decreasing productivity. As a persistent soilborne pathogen causing severe early dying and substantial yield losses, its dominance in the high altitude and cooler areas signals a potential long-term field infestation. These results suggest that further investigation, including soil sampling, tuber health assessments, and pathogenicity testing, would be warranted to clarify the epidemiological significance of Verticillium dahliae in high-altitude sites and to inform potential management decisions. The co-occurrence of multiple Fusarium species, including F. oxysporum and F. solani complex members, across various sub-regions underscores the complexity of potato wilt and dry rot diseases. These combined inoculum occurrences of the Fusarium species can cause increased rotting of the infected tubers, as observed in a study conducted by Tiwari [26] in Australia. The co-occurrence and synergistic interaction between Fusarium and Verticillium, leading to more severe early dying, emphasize the need for integrated management approaches targeting both pathogens. Similarly, the prominent detection of Rhizoctonia solani at Adi Tekeliezan points to localized pressure from stem canker, warranting further characterization of anastomosis groups for tailored management. The identification of Ralstonia solanacearum, a devastating bacterial wilt pathogen, is relatively low but indicates the necessity of continuous surveillance for plants, as even low inoculum can bring about serious disease outbreaks under conducive conditions. This pathogen, responsible for bacterial wilt, is highly destructive and adapted to cool highland environments [27,28].
The alpha diversity analysis revealed that pathogen community complexity varied greatly among locations, where diseased samples were generally more diverse than healthy ones. This could indicate that disease development is dominated by a larger and/or more synergistic pathogen community. The high microbial diversity in diseased plant tissue often reflects complex multi-pathogen infections and secondary colonization rather than a healthy community [29,30]. The distinct clustering of asymptomatic samples with lower diversity supports this, suggesting earlier-stage or simpler early infections. A similar study of microbial community composition associated with potato plants in Canada, analyzed using next-gen sequencing, found differences in alpha diversity of the bacterial communities between the healthy and diseased plants [17].
The high phylogenetic diversity of bacteria observed at specific sites, such as Berik and Serejeka, might suggest the existence of various pathogen lineages due to repeated introductions or local evolution. Recent metagenomic work on potato cyst nematode fields confirms varied bacterial richness tied to host–pathogen dynamics [23].
The contrasting patterns of bacterial and fungal diversity at Logo Anseba (low bacterial, high fungal) and Emni Haili (high bacterial, low fungal) suggest potentially independent drivers of community assembly, the competitive microbial interactions, and distinct disease etiologies, and warrant formal statistical testing in future replicated studies [31,32]. Recent studies confirm that bacterial and fungal diversity often decouple across potato sites due to niche-specific responses, competitive interactions, and disease pressures like potato early dying (PED) due to Verticillium and others. In potato rhizospheres, bacterial communities respond strongly to root proximity and growth stage, while fungal diversity peaks mid-season and is more soil-driven or air-borne, showing distinct abundance patterns (e.g., bacterial shifts in rhizosphere vs. fungal stability in bulk soil). Wilt of potato due to bacterial stress elevates fungal diversity in potato endospheres but suppresses it in rhizospheres, creating inverse profiles akin to low bacterial/high fungal or vice versa scenarios [33,34,35]. Potato monoculture gradients reveal bacteria shaped by cropping, while fungi respond differently, with Fusarium enrichment decoupled from broader bacterial changes [36].
Results of the beta diversity analysis also supported that potato microbial communities, including pathogens, cluster significantly by location due to spatial heterogeneity in highland systems. Although some cluster patterns were associated more with zones of regional sample origin (like Maekel and Debub), the contribution of other parameters, such as microclimatic differences, local agricultural practices (e.g., rotation system), or seed potato multiplication sites, could not be discounted. For example, sites with intensive irrigation or waterlogging may favour the presence of waterborne pathogens, such as Pectobacterium, whereas sites with lower field maintenance or decreased standards of phytosanitary requirements could experience increased populations of soil-borne pathogens. This spatial heterogeneity in potato pathogens aligns with several studies, where rotation, seed quality, cultivar, and microclimate drive variation [1,36,37]. The observed variation in pathogen community composition across sub-regions is consistent with well-documented ecological relationships between altitude, temperature, and plant-associated microbial communities in highland agricultural systems. The dominance of Verticillium dahliae in cooler high-altitude sub-regions aligns with its established preference for lower soil temperatures and well-drained highland soils [38,39]. Similarly, the higher relative abundance of Alternaria-associated taxa in sub-regions with warmer, semi-arid conditions is consistent with published relationships between temperature variability and early blight severity in potato [40]. The geographic clustering of bacterial communities observed in this study parallels patterns reported across altitudinal gradients in comparable highland potato production zones, where altitude and associated climatic gradients are recognized drivers of microbial community structure [33]. Future studies integrating direct environmental measurements, including temperature, humidity, and soil physicochemical parameters, at the farm level will enable formal quantification of the relative contributions of these drivers to pathogen community assembly across Eritrean potato production systems. A limitation of this study is the use of composite, sub-region-level pooled samples, with one pooled sample per sub-region. This was necessitated by logistical constraints of nationwide sampling across 81 farms in 14 sub-regions, and is consistent with published landscape-scale metagenomics surveys employing pooled leaf sampling strategies [9,41]. The pooling reduces farm-level biological resolution, because each sub-region is represented by a single composite sample, formal significance testing of diversity differences was not applicable. The diversity patterns described in this section are therefore descriptive rather than statistically inferred. Future studies should incorporate replicated plant-level sampling.
5. Conclusions
This study has successfully elucidated the genetic diversity and geographic distribution of major bacterial and fungal potato pathogen communities in Eritrea, revealing significant spatial structuring and frequent detection of pathogen-associated taxa, including in asymptomatic plant material. We identified key pathogens such as Alternaria spp., Verticillium dahliae, Fusarium spp., Rhizoctonia solani, and Ralstonia solanacearum, revealing their frequent occurrence across sampled sub-regions with significant regional variation. The consistent detection of pathogen-associated taxa in the asymptomatic plants highlights the potential limitations of visual-only disease surveillance, though viability and pathogenicity confirmation are required. These findings underscore the necessity of moving beyond conventional symptom-based surveillance towards molecular approaches for accurate pathogen detection and monitoring. The baseline data generated here are invaluable for the development of robust, region-specific disease management strategies, a strengthened pathogen surveillance system, enhanced seed quality programmes, and ultimately contribute to improved potato production and food security in Eritrea. Future studies could integrate quantitative PCR (qPCR) to estimate absolute pathogen loads and employ whole-genome sequencing for strain-level differentiation, virulence factors, and evolution tracking, enabling the development of more resilient potato production systems across East African and similar climatic regions.
Supplementary Materials
The following supporting information can be downloaded at: https://www.mdpi.com/article/10.3390/applbiosci5030057/s1, Table S1 Distance Matrix of the Bacterial and Fungal Abundance.
Author Contributions
Conceptualization, M.M., J.O.N., C.M.M. and T.M.; methodology, M.M., J.O.N. and C.M.M.; software, M.M. and S.G.M.; validation, M.M., J.O.N. and C.M.M.; formal analysis, M.M. and S.G.M.; investigation, M.M.; resources, J.O.N., C.M.M. and T.M.; data curation, M.M.; writing—original draft preparation, M.M.; writing—review and editing, M.M., J.O.N., C.M.M., T.M. and S.G.M.; visualization, M.M. and S.G.M.; supervision, J.O.N., C.M.M. and T.M.; project administration, M.M.; funding acquisition, M.M. All authors have read and agreed to the published version of the manuscript.
Funding
This research was funded and supported by the Ministry of Agriculture and Vita under projects EU FOOD/2019/411-806 and EuropeAid/177476/DD/ACT/ER.
Institutional Review Board Statement
Not applicable.
Informed Consent Statement
Not applicable.
Data Availability Statement
The raw sequencing data generated in this study have been deposited in the NCBI Sequence Read Archive (SRA) under BioProject accession number PRJNA1445368. All scripts used for data analysis are available from the corresponding author upon reasonable request.
Acknowledgments
The authors gratefully acknowledge the support of the National Animal and Plant Health Laboratory of Eritrea (NAPHL), the staff of the National Agricultural Research Institute (NARI), Agro-mapping and the Geospatial unit of the Ministry of Agriculture for their support in GIS mapping, and local sub-region extension workers for their invaluable assistance during field sampling. The authors also thank the staff of the Institute of Biotechnology Research (IBR), JKUAT, for laboratory support.
Conflicts of Interest
The authors declare that this study received funding from Vita. Vita supported the logistics and materials needed. Vita is in an Irish developing organization working in Eritrea and currently supporting farmers, specifically in the provision of clean seed potato materials. Vita is registered with the Charities Regulator in Ireland under the Registered Charity Number (RCN) 20024192. The funder had no role in the design of the study; in the collection, analysis, or interpretation of data, in the writing of the manuscript, or in the decision to publish the results.
Abbreviations
The following abbreviations are used in this manuscript:
| ASV | Amplicon Sequence Variant |
| CTAB | Cetyltrimethylammonium bromide |
| DADA2 | Divisive Amplicon Denoising Algorithm 2 |
| ITS | Internal Transcribed Spacer |
| NAPHL | National Animal and Plant Health Laboratory |
| NARI | National Agricultural Research Institute |
| NCBI | National Centre for Biotechnology Information |
| PAC | Powdered Activated Charcoal |
| PBS | Phosphate-Buffered Saline |
| PCoA | Principal Coordinates Analysis |
| PD | Phylogenetic Diversity |
| QIIME | Quantitative Insights Into Microbial Ecology |
| rRNA | Ribosomal Ribonucleic Acid |
| SRA | Sequence Read Archive |
| SDS | Sodium Dodecyl Sulfate |
| UPGMA | Unweighted Pair Group Method with Arithmetic Mean |
References
- Harahagazwe, D.; Condori, B.; Barreda, C.; Bararyenya, A.; Byarugaba, A.A.; Kude, D.A.; Lung’Aho, C.; Martinho, C.; Mbiri, D.; Nasona, B.; et al. How Big Is the Potato (Solanum tuberosum L.) Yield Gap in Sub-Saharan Africa and Why? A Participatory Approach. Open Agric. 2018, 3, 180–189. [Google Scholar] [CrossRef] [Scilit]
- Villamor, D.E.V.; Ho, T.; Al Rwahnih, M.; Martin, R.R.; Tzanetakis, I.E. High Throughput Sequencing for Plant Virus Detection and Discovery. Am. Phytopathol. Soc. 2019, 109, 716–725. [Google Scholar] [CrossRef] [Scilit]
- Della Bartola, M.; Byrne, S.; Mullins, E. Characterization of Potato Virus Y Isolates and Assessment of Nanopore Sequencing to Detect and Genotype Potato Viruses. Viruses 2020, 12, 478. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Roossinck, M.J. Deep Sequencing for Discovery and Evolutionary Analysis of Plant Viruses. Virus Res. 2017, 239, 82–86. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Massart, S.; Olmos, A.; Jijakli, H.; Candresse, T. Current Impact and Future Directions of High Throughput Sequencing in Plant Virus Diagnostics. Virus Res. 2014, 188, 90–96. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Walters, W.; Hyde, E.R.; Berg-Lyons, D.; Ackermann, G.; Humphrey, G.; Parada, A.; Gilbert, J.A.; Jansson, J.K.; Caporaso, J.G.; Fuhrman, J.A.; et al. Improved Bacterial 16S RRNA Gene (V4 and V4-5) and Fungal Internal Transcribed Spacer Marker Gene Primers for Microbial Community Surveys. mSystems 2015, 1, e00009-15. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Muñoz-Ramírez, Z.Y.; González-Escobedo, R.; Avila-Quezada, G.D.; Ramírez-Sánchez, O.; Higareda-Alvear, V.M.; Zapata-Chávez, E.; Borrego-Loya, A.; Muñoz-Castellanos, L.N. Exploring Microbial Rhizosphere Communities in Asymptomatic and Symptomatic Apple Trees Using Amplicon Sequencing and Shotgun Metagenomics. Agronomy 2024, 14, 357. [Google Scholar] [CrossRef] [Scilit]
- Yu, X.L.; Hu, X.Y.; Wang, X.X.; Zhang, X.Y.; Du, K.B. A Protocol Specialized for Microbial DNA Extraction from Living Poplar Wood. Not. Bot. Horti Agrobot. Cluj. Napoca. 2022, 50, 12822. [Google Scholar] [CrossRef] [Scilit]
- Orn, B.; Lindahl, D.; Nilsson, R.H.; Tedersoo, L.; Abarenkov, K.; Carlsen, T.; Kjøller, R.; Kõljalg, U.; Pennanen, T.; Rosendahl, S.; et al. Methods Fungal Community Analysis by High-Throughput Sequencing of Amplified Markers-a User’s Guide. New Phytol. 2013, 199, 288–299. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Martin, M. Cutadapt Removes Adapter Sequences from High-Throughput Sequencing Reads. EMBnet. J. 2011, 17, 10–12. [Google Scholar] [CrossRef] [Scilit]
- Callahan, B.J.; McMurdie, P.J.; Rosen, M.J.; Han, A.W.; Johnson, A.J.A.; Holmes, S.P. DADA2: High-Resolution Sample Inference from Illumina Amplicon Data. Nat. Methods 2016, 13, 581–583. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Bolyen, E.; Rideout, J.R.; Dillon, M.R.; Bokulich, N.A.; Abnet, C.C.; Al-Ghalith, G.A.; Alexander, H.; Alm, E.J.; Arumugam, M.; Asnicar, F.; et al. Reproducible, interactive, scalable and extensible microbiome data science using QIIME 2. Nat. Biotechnol. 2019, 37, 852–857. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Gildemacher, P.R.; Kaguongo, W.; Ortiz, O.; Tesfaye, A.; Woldegiorgis, G.; Wagoire, W.W.; Kakuhenzire, R.; Kinyae, P.M.; Nyongesa, M.; Struik, P.C.; et al. Improving Potato Production in Kenya, Uganda and Ethiopia: A System Diagnosis. Potato Res. 2009, 52, 173–205. [Google Scholar] [CrossRef] [Scilit]
- Onditi, J.; Nyongesa, M.; Vlugt, R. Prevalence, Distribution and Control of Six Major Potato Viruses in Kenya. Trop. Plant Pathol. 2021, 46, 311–323. [Google Scholar] [CrossRef] [Scilit]
- Guchi, E. Disease Management Practice on Potato (Solanum tuberosum L.) in Ethiopia. World J. Agric. Res. 2015, 3, 34–42. [Google Scholar]
- Aleti, G.; Nikolić, B.; Brader, G.; Pandey, R.V.; Antonielli, L.; Pfeiffer, S.; Oswald, A.; Sessitsch, A. Secondary Metabolite Genes Encoded by Potato Rhizosphere Microbiomes in the Andean Highlands Are Diverse and Vary with Sampling Site and Vegetation Stage. Sci. Rep. 2017, 7, 2330. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Borza, T.; Lumactud, R.A.; Shim, S.Y.; Al-Mughrabi, K.; Prithiviraj, B. Microbial Community Composition Associated with Potato Plants Displaying Early Dying Syndrome. Microorganisms 2025, 13, 1482. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Meno, L.; Abuley, I.; Seijo, M.C.; Escuredo, O. Management Strategies for Early Blight in Potatoes: Assessment of the TOMCAST Model, Including the Aerobiological Risk Level and Critical Phenological Period. Agriculture 2024, 14, 1414. [Google Scholar] [CrossRef] [Scilit]
- da Silva Silveira Duarte, H.; Zambolim, L.; Machado, F.J.; Pereira Porto, H.R.; Rodrigues, F.Á. Comparative Epidemiology of Late Blight and Early Blight of Potato under Different Environmental Conditions and Fungicide Application Programs. Semin. Cienc. Agrar. 2019, 40, 1805–1818. [Google Scholar] [CrossRef] [Scilit]
- Davies, T.J.; Pedersen, A.B. Phylogeny and Geography Predict Pathogen Community Similarity in Wild Primates and Humans. Proc. R. Soc. B Biol. Sci. 2008, 275, 1695. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Chaloner, T.M.; Gurr, S.J.; Bebber, D.P. Plant Pathogen Infection Risk Tracks Global Crop Yields under Climate Change. Nat. Clim. Change 2021, 11, 710–715. [Google Scholar] [CrossRef] [Scilit]
- Bebber, D.P.; Gurr, S.J. Crop-Destroying Fungal and Oomycete Pathogens Challenge Food Security. Fungal Genet. Biol. 2015, 74, 62–64. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Kiige, J.K.; Kavoo, A.M.; Mwajita, M.R.; Mogire, D.; Ogada, S.; Wekesa, T.B.; Kiirika, L.M. Metagenomic Characterization of Bacterial Abundance and Diversity in Potato Cyst Nematode Suppressive and Conducive Potato Rhizosphere. PLoS ONE 2025, 20, e0323382. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Zhang, X.; Zhang, X. Genetic and Biological Characterization of Bacterial Complexes Causing Potato Blackleg and Soft Rot. Ph.D. Thesis, University of Maine, Orono, ME, USA, 2025. [Google Scholar]
- Hameed, A.; Zeeshan, M.; Binyamin, R.; Alam, M.W.; Ali, S.; Zaheer, M.S.; Ali, H.; Riaz, M.W.; Ali, H.H.; Elshikh, M.S.; et al. Molecular Characterization of Pectobacterium atrosepticum Infecting Potato and Its Management through Chemicals. PeerJ 2024, 12, e17518. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Tiwari, R.K.; Lal, M.K.; Kumar, R.; Sharma, S.; Sagar, V.; Kumar, A.; Singh, B.; Aggarwal, R. Impact of Fusarium Infection on Potato Quality, Starch Digestibility, In Vitro Glycemic Response, and Resistant Starch Content. J. Fungi 2023, 9, 466. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Charkowski, A.; Sharma, K.; Parker, M.L.; Secor, G.A.; Elphinstone, J. Bacterial Diseases of Potato. In The Potato Crop; Springer International Publishing: Cham, Switzerland, 2020; pp. 351–388. [Google Scholar]
- Wang, Z.; Luo, W.; Cheng, S.; Zhang, H.; Zong, J.; Zhang, Z. Ralstonia Solanacearum—A Soil Borne Hidden Enemy of Plants: Research Development in Management Strategies, Their Action Mechanism and Challenges. Front. Plant Sci. 2023, 14, 1141902. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Compant, S.; Samad, A.; Faist, H.; Sessitsch, A. A Review on the Plant Microbiome: Ecology, Functions, and Emerging Trends in Microbial Application. J. Adv. Res. 2019, 19, 29–37. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Trivedi, P.; Mattupalli, C.; Eversole, K.; Leach, J.E. Enabling Sustainable Agriculture through Understanding and Enhancement of Microbiomes. New Phytol. 2021, 230, 2129–2147. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Zimudzi, J.; van der Waals, J.E.; Coutinho, T.A.; Cowan, D.A.; Valverde, A. Temporal Shifts of Fungal Communities in the Rhizosphere and on Tubers in Potato Fields. Fungal Biol. 2018, 122, 928–934. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Hardoim, P.R.; van Overbeek, L.S.; Berg, G.; Pirttilä, A.M.; Compant, S.; Campisano, A.; Döring, M.; Sessitsch, A. The Hidden World within Plants: Ecological and Evolutionary Considerations for Defining Functioning of Microbial Endophytes. Microbiol. Mol. Biol. Rev. 2015, 79, 293–320. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Bak, G.R.; Lee, K.K.; Clark, I.M.; Mauchline, T.H.; Kavamura, V.N.; Jee, S.; Lee, J.T.; Kim, H.; Lee, Y.H. Changes in the Potato Rhizosphere Microbiota Richness and Diversity Occur in a Growth Stage-Dependent Manner. Sci. Rep. 2025, 15, 2284. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Liu, X.; Zhang, J.; Gu, T.; Zhang, W.; Shen, Q.; Yin, S.; Qiu, H. Microbial Community Diversities and Taxa Abundances in Soils along a Seven-Year Gradient of Potato Monoculture Using High Throughput Pyrosequencing Approach. PLoS ONE 2014, 9, e86610. [Google Scholar] [CrossRef] [Scilit]
- Li, Z.; Zhu, J.; Shi, T.; Gao, C.; Qiu, X.; Bao, M.; Li, Y.; Bi, Z.; Yao, P.; Sun, C.; et al. Microbial Community Restructuring under Crop Rotation: A Sustainable Strategy to Counteract Potato Monoculture-Induced Soil Degradation in Arid Ecosystems. Agric. Ecosyst. Environ. 2026, 400, 110245. [Google Scholar] [CrossRef] [Scilit]
- Charkowski, A.O. The Changing Face of Bacterial Soft-Rot Diseases. Annu. Rev. Phytopathol. 2018, 56, 269–288. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Imam, N.; Belda, I.; García-Jiménez, B.; Duehl, A.J.; Doroghazi, J.R.; Almonacid, D.E.; Thomas, V.P.; Acedo, A. Local Network Properties of Soil and Rhizosphere Microbial Communities in Potato Plantations Treated with a Biological Product Are Important Predictors of Crop Yield. mSphere 2021, 6, e0013021. [Google Scholar] [CrossRef] [Scilit]
- Klosterman, S.J.; Atallah, Z.K.; Vallad, G.E.; Subbarao, K.V. Diversity, Pathogenicity, and Management of Verticillium Species. Annu. Rev. Phytopathol. 2009, 47, 39–62. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Wheeler, D.L.; Dung, J.K.S.; Johnson, D.A. From Pathogen to Endophyte: An Endophytic Population of Verticillium dahliae Evolved from a Sympatric Pathogenic Population. New Phytol. 2019, 222, 497–510. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Escuredo, O.; Seijo-Rodríguez, A.; Meno, L.; Rodríguez-Flores, M.S.; Seijo, M.C. Seasonal Dynamics of Alternaria during the Potato Growing Cycle and the Influence of Weather on the Early Blight Disease in North-West Spain. Am. J. Potato Res. 2019, 96, 532–540. [Google Scholar] [CrossRef] [Scilit]
- Vidal-Villarejo, M.; Hammerschmidt, M.; Oppliger, B.; Oppliger, H.; Schmid, K. Exotic Genetic Lineages Rapidly Replace Established Exserohilum turcicum Populations in the Swiss Rhine Valley. Plant Pathol. 2026, 75, e70144. [Google Scholar] [CrossRef] [Scilit]

Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2026 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license.