


Abstract
CO2 adsorption on subnanometric metal clusters is highly sensitive to the computational protocol used to describe the potential energy surface, particularly when several low-lying geometries and spin states are accessible. In this work, CO2 adsorption on Cu4 and Sc4 clusters was investigated using density functional theory (DFT) to evaluate how the choice of functional/basis-set protocol, spin multiplicity, initial geometry, and vibrational stability affects the predicted adsorption behavior. Four representative computational protocols (TPSSh, r2SCAN-3c, PBE-D4/def2-TZVP, and PBE0-SDD) were assessed for isolated clusters and cluster–CO2 complexes. The lowest harmonic vibrational frequency, ωmin, was used as a diagnostic criterion to distinguish true minima from unstable or weakly defined stationary points. Selected cases were also cross-checked using the ORCA and Gaussian quantum-chemistry packages to assess whether comparable computational settings yielded consistent stationary-point character. The results show that Cu4 generally exhibits weak CO2 binding, whereas Sc4 displays stronger but more protocol-dependent adsorption, consistent with its higher structural flexibility and more pronounced Lewis-acid character. Low-frequency and imaginary modes were found in several optimized structures, indicating that adsorption energies should not be interpreted without prior vibrational validation. The comparison also shows that variations in functional/basis-set treatment and spin multiplicity can alter both the optimized geometry and the predicted adsorption strength. Therefore, CO2 adsorption on small metal clusters should be discussed using combined structural, vibrational, and energetic criteria rather than electronic adsorption energies alone. Overall, this study provides a protocol-oriented framework for evaluating the reliability of DFT predictions in CO2 adsorption on Cu4 and Sc4 clusters.
1. Introduction
CO2 capture and utilization are central topics in materials chemistry and catalysis because they address both emission mitigation and the transformation of CO2 into value-added carbon feedstocks [1,2,3]. Beyond its environmental relevance, CO2 conversion is directly connected with the development of circular chemical processes in which carbon-containing waste streams can be reintegrated into productive routes [4,5,6,7]. At the molecular level, the first step in many capture and activation processes is the interaction of CO2 with a metallic site. This interaction controls the adsorption strength and may induce structural perturbations in the molecule, including variation in the O–C–O angle and elongation of the C–O bonds [8,9,10]. These geometric changes are commonly interpreted as early indicators of CO2 activation. However, their magnitude and chemical meaning depend strongly on the nature of the metal center and on the theoretical description used to model the system. In metal–CO2 interactions, electrostatics, polarization, charge transfer, metal-to-CO2 back-donation into π* orbitals, and dispersion forces can contribute simultaneously. Therefore, the predicted adsorption mode and adsorption energy are not purely structural outputs, but are directly affected by the computational protocol [11].
Small metal clusters provide molecular-scale models of undercoordinated active sites and can bridge the gap between isolated coordination complexes and extended metallic surfaces. Their low nuclearity makes them computationally accessible, while their electronic structure remains sufficiently complex to exhibit several relevant phenomena, including multiple low-lying isomers, soft vibrational modes, and closely spaced spin states [12,13,14]. In this context, Cu4 and Sc4 were selected because they represent two chemically contrasting cases for CO2 adsorption. Copper clusters are expected to show relatively weak and often reversible interactions with CO2 due to the more filled character of the Cu d manifold and the limited tendency of Cu to polarize the adsorbate strongly. In contrast, scandium clusters are more electropositive and can behave as stronger Lewis-acidic sites, favoring CO2 polarization through oxygen coordination and stronger metal–adsorbate electrostatic interactions. Therefore, the Cu4/Sc4 comparison is not arbitrary; it allows the evaluation of two distinct adsorption regimes: weak adsorption dominated by modest polarization in Cu4 and stronger, more structurally adaptive adsorption in Sc4.
Tetrameric clusters were chosen as minimal but chemically meaningful models. Compared with dimers or trimers, M4 clusters can support planar and three-dimensional arrangements, symmetry changes, and spin-dependent structural reorganization. At the same time, they remain small enough to permit systematic testing of several density functional theory (DFT) protocols, spin multiplicities, and initial geometries. This balance is important because subnanometric clusters often have relatively flat potential energy surfaces, where small changes in the exchange–correlation functional, basis set, effective core potential (ECP), dispersion correction, integration grid, convergence criteria, or optimization algorithm can lead to different stationary points [15]. Consequently, a single optimized structure or a single electronic adsorption energy may not be sufficient to define a reliable adsorption trend.
From a methodological perspective, benchmarking is necessary to avoid overinterpreting results arising from a particular functional/basis-set combination rather than from the intrinsic chemistry of the cluster–CO2 system [16,17]. This is especially relevant for small transition-metal clusters, where geometry, spin state, and vibrational stability are strongly coupled. In this work, the term “computational protocol” refers to the combined use of the exchange–correlation functional, basis set, ECP when applicable, dispersion treatment, and numerical settings. Four representative protocols—TPSSh, r2SCAN-3c, PBE-D4/def2-TZVP, and PBE0-SDD—were evaluated to assess their influence on the optimized geometries and adsorption energies of Cu4–CO2 and Sc4–CO2 complexes. The use of r2SCAN-3c provides a compact composite DFT approach, PBE-D4/def2-TZVP allows evaluation of a generalized gradient approximation with explicit dispersion and a triple-ζ basis set, TPSSh provides a meta-hybrid reference commonly used in transition-metal chemistry, and PBE0-SDD allows the effect of a hybrid functional combined with an ECP-based description of the metal centers to be assessed [18,19,20].
The present study aims to evaluate the sensitivity of CO2 adsorption on Cu4 and Sc4 clusters to the choice of DFT protocol, spin multiplicity, initial geometry, and vibrational stability. The lowest harmonic frequency, ωmin, was used as an operational descriptor to distinguish true minima from unstable stationary points or shallow regions of the potential energy surface. In addition, selected calculations were compared between the ORCA and Gaussian quantum-chemistry packages. This comparison was not intended as a general benchmark of software performance, but as a limited cross-validation of whether comparable computational settings produce consistent stationary-point character. To support the methodological assessment, Cu2 was used as an auxiliary reference system through experimentally comparable observables, including r(Cu–Cu), vibrational frequency, ionization potential, and electron affinity. Overall, this work seeks to establish a reliability-oriented framework for interpreting CO2 adsorption on small metal clusters, emphasizing that adsorption energies should be evaluated together with geometric descriptors, vibrational validation, and protocol-dependent uncertainty [21].
2. Results and Discussion
2.1. Geometric Reorganization of Cu4 After Optimization
The initial and optimized Cu4 structures were compared for three starting symmetries and two spin multiplicities to evaluate structural reorganization and protocol-dependent geometric sensitivity, as summarized in Figure 1.
Figure 1.
Initial and optimized structures of Cu4 clusters with D2h, D3h, and Td starting symmetries, considering singlet and triplet multiplicities across the evaluated DFT protocols.
Figure 1 indicates that Cu4 cannot be treated as a rigid tetrameric unit, even before CO2 adsorption. The optimized geometry depends simultaneously on the initial symmetry, spin multiplicity, and computational protocol, confirming that the Cu4 potential energy surface contains several closely related structural basins. For the D2h starting geometry, the rhombohedral framework is generally preserved after optimization; however, the magnitude of the Cu–Cu contraction or elongation varies across the evaluated methods. This behavior indicates that each protocol describes the balance between metal–metal bonding, exchange–correlation effects, and spin-dependent electron redistribution differently.
In contrast, Figure 1 shows that the D3h starting geometry is more likely to lose its initial symmetry during optimization. Several optimized structures evolve toward lower-symmetry arrangements, indicating that not all protocols stabilize the D3h motif equally. This structural instability is consistent with a shallow region of the potential energy surface, where small changes in the functional treatment or spin state can redirect the optimization toward neighboring minima. Therefore, adsorption energies obtained from D3h-derived structures must be interpreted cautiously, because differences in Eads may arise not only from the metal–CO2 interaction itself, but also from changes in the underlying Cu4 isomer [12,22,23].
For the Td starting geometry, the three-dimensional character is more clearly retained, although method-dependent distortions remain. This result is relevant because three-dimensional Cu4 motifs may expose coordination environments for CO2 adsorption that differ from those available in planar geometries. Overall, Figure 1 establishes the structural basis for the subsequent adsorption analysis. Before comparing Eads values, it is necessary to verify whether each protocol preserves the same cluster topology or drives the system toward a different optimized minimum. This justifies the later use of vibrational validation and stationary-point analysis as mandatory filters for reliable interpretation of CO2 adsorption on Cu4 [19,22,24].
2.2. Geometric Metrics for Cu4 D2h and Method and Spin Effect
The D2h Cu4 structures were analyzed to determine whether the optimized metal framework preserves the bond equivalence of the initial geometry or undergoes protocol-dependent contraction. This comparison, summarized in Table 1, is necessary before interpreting CO2 adsorption because changes in the isolated cluster geometry modify the electronic structure and the local coordination environment available for adsorbate binding.
Table 1.
Cu–Cu bond distances for the initial and optimized D2h Cu4 clusters calculated using different DFT protocols and spin multiplicities.
As shown in Table 1, the optimized D2h Cu4 structures exhibit a clear dependence on both the computational protocol and spin multiplicity. The initial geometry contains four equivalent Cu–Cu distances of 2.40348 Å, whereas geometry optimization produces method-dependent contraction or slight elongation of the metallic framework. In the singlet state, PBE-D4 and TPSSh preserve relatively long Cu–Cu distances of ~2.41 Å, indicating a less compact metal framework. In contrast, r2SCAN-3c and PBE0-SDD predict shorter Cu–Cu contacts. This contraction becomes more pronounced under triplet conditions, where selected Cu–Cu distances decrease to approximately 2.31–2.33 Å.
This trend indicates that the description of Cu–Cu bonding is sensitive to the exchange–correlation functional, basis/ECP treatment, and spin configuration. The stronger contraction observed for the triplet structures suggests that spin-state changes can modify the distribution of electron density within the Cu4 framework and, consequently, the availability of frontier metal states involved in CO2 polarization and metal-to-CO2 donation/back-donation interactions. Therefore, Table 1 provides more than a geometric comparison; it establishes the structural reference required to interpret subsequent adsorption energies. If the isolated cluster is described with different degrees of compactness depending on the method, variations in Eads may reflect not only the metal–CO2 interaction, but also differences in the initial electronic structure and relaxation capacity of the Cu4 cluster [10,19,25,26].
2.3. Geometric Instability of Cu4 D3h and Loss of Symmetry
The D3h Cu4 topology was examined to evaluate whether this higher-symmetry arrangement corresponds to a stable structural motif or to a shallow region of the potential energy surface that relaxes toward lower-symmetry configurations. The corresponding Cu–Cu structural descriptors are summarized in Table 2.
Table 2.
Cu–Cu structural descriptors for the initial and optimized D3h Cu4 clusters calculated using different DFT protocols and spin multiplicities.
As shown in Table 2, the D3h arrangement is not consistently preserved after geometry optimization. Several optimized structures exhibit asymmetric Cu–Cu distances or missing equivalent contacts, indicating that the initial D3h motif relaxes toward lower-symmetry geometries depending on the computational protocol and spin state. This behavior suggests that D3h Cu4 lies in a shallow region of the potential energy surface, where small changes in exchange–correlation treatment, basis-set representation, or spin configuration can drive the system toward different local minima.
This result is important for the adsorption analysis because, as indicated by Table 2, Eads values obtained across different final cluster geometries cannot be interpreted solely in terms of electronic differences. They may also include contributions from isomeric reorganization. Therefore, Table 2 provides a structural filter for assessing whether adsorption energies are being compared for equivalent or non-equivalent Cu4 frameworks [12,22,25].
2.4. Cross-Validation Between Codes Using ωmin as Minimum True Control
To evaluate whether the optimized Cu4 structures correspond to true minima or unstable stationary points, the lowest harmonic frequency, ωmin, was compared between selected Gaussian reference data and ORCA calculations. The comparison, summarized in Table 3, focuses on the final symmetry, relative stationary-point character, and presence or absence of imaginary frequencies.
Table 3.
Inter-code comparison of Cu4 stationary-point character using ωmin and final symmetry as vibrational-stability descriptors.
As shown in Table 3, nominally comparable computational protocols do not always lead to the same stationary-point character. For the D2h singlet, both Gaussian and ORCA produce positive ωmin values close to 60 cm−1, supporting the assignment of this structure as a reproducible true minimum. In contrast, for the D2h triplet, the Gaussian reference reports an imaginary frequency, whereas the ORCA calculation gives a positive ωmin. This difference was interpreted from the sign of the lowest harmonic frequency and the final optimized geometry obtained with each code. Within the harmonic approximation, an imaginary frequency indicates negative curvature along at least one normal coordinate, while a positive ωmin indicates the absence of negative curvature in the local Hessian [27,28].
Shows that this behavior is not restricted to one geometry. For D3h-derived structures, changes in final symmetry and ωmin indicate that the optimization pathway may lead to different local regions of the potential energy surface depending on the code implementation. Similarly, Td-derived structures include cases with imaginary frequencies and cases with positive ωmin, confirming that the Cu4 potential energy surface contains soft distortion coordinates and nearby stationary points.
Table 3 should not be interpreted as a simple comparison of total energies. Its main purpose is to ensure reproducibility and vibrational stability before using optimized structures in the adsorption-energy analysis. When nominally similar calculations produce different ωmin signs or different final symmetries, the corresponding Eads values must be interpreted cautiously, because they may reflect differences in the stationary-point character rather than only differences in the metal–CO2 interaction strength.
2.5. Ωmin Statistics and Quantitative Criterion of Vibrational Stability
The distribution of ωmin values was summarized by separating stable and unstable cases to quantify the frequency with which real and imaginary frequencies occur in the benchmark. The resulting statistical descriptors are presented in Table 4.
Table 4.
Statistical summary of ωmin values grouped according to real frequencies (ω > 0) and imaginary frequencies (ω < 0).
As shown in Table 4, the ωmin distribution reflects a characteristic feature of small metal clusters: the presence of structurally soft stationary points with low-frequency vibrational modes. The group of real frequencies (ω > 0) shows a mean value close to 48 cm−1, with moderate dispersion, indicating that even the verified minima are associated with relatively shallow potential-energy surfaces. These results are consistent with true minima that remain vibrationally soft and therefore sensitive to small structural distortions.
In contrast, the group of imaginary frequencies (ω < 0) has a mean value of approximately −42 cm−1 and a broader dispersion, with a minimum as negative as −88.92 cm−1. As indicated by Table 4, these values are not compatible with merely negligible numerical noise, but instead suggest the presence of genuine instabilities associated with low-barrier distortion coordinates. Such coordinates may involve out-of-plane deformations, incipient isomerization, or structural relaxations coupled to electronic reorganization [29,30].
Table 4 supports the use of ωmin as an operational quality descriptor rather than as secondary vibrational information. By statistically separating real from imaginary frequencies, the table defines which optimized structures can be treated as genuinely comparable minima for discussing CO2 adsorption and activation, and which should be excluded from direct energetic interpretation.
2.6. Protocol Validation with Cu2 Using IP and EA as Sensitive Observables
The electronic properties of the Cu2 dimer were compared with experimental reference values to provide an external validation criterion for selecting a computational protocol with lower risk of methodological artifacts, as shown in Figure 2.
Figure 2.
Comparison of the ionization potential (IP) and electron affinity (EA) of the Cu2 dimer calculated using the evaluated DFT methods against experimental reference values [31].
As shown in Figure 2, this comparison allows evaluation of whether each method reproduces the energy differences between charged states, which are highly sensitive to exchange–correlation treatment, self-interaction error, basis-set quality, and ECP treatment. The IP predicted by r2SCAN-3c is close to the experimental value, and TPSSh also shows good agreement, suggesting that both methods reasonably describe the HOMO energy and the balance of stabilization upon ionization.
For EA, Figure 2 shows that PBE-D4 approaches the experimental value, whereas r2SCAN-3c underestimates it, and PBE0-SDD even gives a negative EA. This behavior indicates a poor description of the anionic state, likely due to limited diffuse character in the basis set, ECP imbalance, or an excessive fraction of exact exchange for the available basis-space description [31]. The useful conclusion for the benchmark is that r2SCAN-3c is not necessarily the best method for each isolated observable, but it maintains global coherence without catastrophic failures. For this reason, it is defensible as a practical reference protocol for studying adsorption, where metal–CO2 charge-transfer effects are central.
2.7. Geometric Validation of Cu2 Using Cu–Cu Distance
The Cu–Cu equilibrium distance of the Cu2 dimer was compared with the experimental reference value to evaluate how each protocol describes the balance of the metal–metal bond, as summarized in Figure 3.
Figure 3.
Cu–Cu bond distance in the Cu2 dimer calculated using the evaluated DFT methods and compared with the experimental reference value [31].
As shown in Figure 3, r2SCAN-3c reproduces the experimental Cu–Cu distance very closely, while PBE0-SDD also provides a competitive description of the metal–metal bond. In contrast, TPSSh and PBE-D4 predict longer Cu–Cu distances, indicating a slight expansion of the dimer relative to the experimental reference [31]. This elongation suggests that these protocols describe a less compact metal–metal interaction, probably due to differences in the exchange–correlation balance, basis-set treatment, or effective repulsion in the Cu–Cu bonding region.
This geometric validation is relevant for the Cu4 adsorption study because the electron-density distribution and polarizability of the metallic framework influence CO2 binding and possible metal-to-CO2 donation/back-donation interactions. A method that already overestimates the Cu–Cu distance in Cu2 may alter the electronic structure used to describe CO2 adsorption in larger Cu clusters. Therefore, Figure 3 supports the use of r2SCAN-3c as a practical reference protocol for subsequent adsorption analysis, while also emphasizing that geometric accuracy should be considered alongside vibrational and energetic criteria.
2.8. Vibrational Validation of Cu2 Using Fundamental Frequency
The Cu–Cu stretching frequency of the Cu2 dimer was compared with the experimental reference value to evaluate how each protocol describes the local curvature and stiffness of the metal–metal bond potential, as summarized in Figure 4.
Figure 4.
Fundamental Cu–Cu stretching frequency of the Cu2 dimer calculated using the evaluated DFT methods and compared with the experimental reference value [31].
As shown in Figure 4, PBE0-SDD overestimates the experimental Cu–Cu stretching frequency, indicating an excessively rigid potential near the equilibrium geometry [31]. In contrast, TPSSh and PBE-D4 underestimate this frequency, suggesting a softer potential and a less stiff description of the metal–metal interaction. In this comparison, r2SCAN-3c yields an intermediate result and more closely reproduces the experimental vibrational curvature, consistent with its balanced performance across structural and energetic properties.
This result is relevant for the adsorption study because low-frequency modes and soft potential-energy surfaces play a central role in the structural rearrangement of small metal clusters during CO2 binding. A protocol that stiffens the Cu–Cu interaction excessively may overestimate distortion barriers and overstabilize specific optimized geometries, whereas an overly soft description may exaggerate structural reorganization. Therefore, the vibrational agreement shown in Figure 4 supports the use of r2SCAN-3c as a practical reference for the Cu4 and Sc4 adsorption analysis, without introducing extreme rigidity or softness biases in the metal framework.
2.9. Structural Definition of CO2 Adsorption Modes on Cu
The CO2 coordination modes considered in the adsorption study are introduced in Figure 5, and these motifs are subsequently used to interpret differences in optimized geometry and adsorption energy.
Figure 5.
CO2 adsorption modes considered for Cu4 clusters, including η1-C, μ2-O, μ3, and μ4.
As defined in Figure 5, the main CO2 adsorption motifs considered for the Cu4 clusters are not only structural labels, but also distinct interaction scenarios between the adsorbate and the metallic framework. In the η1-C mode, the carbon atom of CO2 interacts directly with a Cu center, which may favor polarization of the C–O bonds and possible metal-to-CO2 donation/back-donation contributions involving the π* orbitals of CO2. In the μ2-O mode, one oxygen atom coordinates to the metallic framework, reflecting a more Lewis acid–base type interaction. In turn, the μ3 and μ4 arrangements involve a larger number of Cu atoms interacting with the adsorbate, which may distribute the metal–CO2 interaction over several centers and promote stronger geometric distortion of the molecule.
Therefore, Figure 5 provides the structural basis for interpreting the optimized Cu4–CO2 complexes discussed in the following section. It also serves as a reference for distinguishing weak molecular adsorption, in which CO2 remains only slightly perturbed, from configurations in which CO2 may undergo stronger geometric and electronic activation through bending of the O–C–O angle, elongation of the C–O bonds, or charge transfer from the metallic cluster.
2.10. Optimized Cu4 CO2 Geometries and Dependence on Symmetry and Mode
The optimized structures of the Cu4–CO2 complexes were analyzed to relate the adsorption geometry to the type of coordination mode and its possible influence on the relative stability of the adsorbed configurations, as summarized in Figure 6.
Figure 6.
Optimized geometries of CO2 adsorption on Cu4 clusters obtained using different initial symmetries, spin multiplicities, and computational methods. Structures 1–12 correspond to singlet Cu4–CO2 systems, whereas structures 25–32 correspond to triplet Cu4–CO2 systems. Atom colors: Cu, cyan; C, yellow; O, red.
As shown in Figure 6, Cu4 exhibits several coordination patterns for CO2 adsorption, even within the same initial symmetry. In several D2h-derived structures, bridge-type configurations are observed in which the carbon and oxygen atoms interact with different Cu centers. This type of arrangement is usually associated with greater stabilization through cooperative coupling and with a higher probability of O–C–O bending. In other D2h cases, oxygen binds more locally to a single Cu atom, corresponding to a more direct coordination mode with greater rotational freedom. Such configurations may be more enthalpically stable in some protocols, but not necessarily more activating, because charge transfer toward the C–O axis may be reduced if the carbon atom is not simultaneously involved in the interaction [32].
Figure 6 also shows that the Td-derived structures display variations in inclination and cases of minimal interaction, indicating that the three-dimensional geometry may present less accessible adsorption sites or an electronic distribution that does not favor CO2 anchoring under certain protocols. This observation prepares the energetic interpretation, because if a method favors bridge-type geometries, it may also predict more negative Eads values. At the same time, such stabilization may increase the risk of overadsorption when dispersion contributions dominate or when the protocol underestimates short-range repulsion.
As shown in Table 5, the charge-transfer response of Cu4–CO2 singlet complexes depends strongly on both the initial symmetry and the computational protocol. For the D2h and D3h arrangements, r2SCAN-3c predicts substantial electron accumulation on CO2, with qCO2 values close to −0.62 e. In contrast, PBE-D4/def2-TZVP and TPSSh/def2-TZVP give qCO2 values close to zero for the same planar arrangements, indicating weak adsorption with limited electronic activation. Therefore, in these cases, favorable or near-thermoneutral Eads values should not be interpreted automatically as evidence of CO2 activation.
Table 5.
NPA charge-transfer analysis for CO2 adsorption on Cu4 singlet complexes. qCO2 was calculated as the sum of the NPA charges of the atoms belonging to the adsorbed CO2 molecule, qCO2 = qC + qO1 + qO2, whereas qCu4 corresponds to the net charge of the metallic fragment. Negative qCO2 values indicate net electron accumulation on CO2, while values close to zero indicate negligible charge transfer. Rows affected by ECP/NPA charge-convention inconsistencies were retained only as qualitative records and were not used for direct quantitative comparison.
The Td singlet configurations in Table 5 show a more pronounced electronic response. Both r2SCAN-3c and PBE-D4/def2-TZVP predict clearly negative qCO2 values, indicating significant metal-to-CO2 charge transfer. This trend is consistent with the more negative adsorption energies obtained for Td-derived Cu4 complexes and suggests that the three-dimensional arrangement provides a more favorable electronic environment for CO2 activation. However, TPSSh/def2-TZVP predicts a negative Eads for Td Cu4 while qCO2 remains close to zero. This discrepancy indicates that the stabilization predicted by TPSSh is not primarily due to net charge transfer to CO2. Still, it may instead arise from geometry-dependent polarization or other method-dependent energetic contributions [25,33,34].
2.11. Sc4 Structural Reorganization in D2h and Expansion of the Metal Network
The initial and optimized geometries of Sc4 with D2h symmetry were compared to assess changes in network size and structural flexibility relative to those of Cu4, as summarized in Figure 7.
Figure 7.
Initial and optimized structures of Sc4 clusters with D2h symmetry, considering singlet and triplet multiplicities across the evaluated DFT protocols.
Figure 7 evidences a marked expansion of the Sc–Sc framework after optimization of the D2h Sc4 structures, in agreement with the larger metal–metal distances later quantified by the corresponding structural descriptors. This expansion suggests weaker Sc–Sc bonding and a more flexible metallic network than that observed for Cu4. Such behavior is chemically consistent with the greater electropositivity of scandium and its lower tendency to form compact metal–metal frameworks.
This structural flexibility is relevant for CO2 adsorption because a more open Sc4 framework may facilitate the formation of accessible binding environments and accommodate multidentate coordination patterns. However, the same flexibility also increases methodological sensitivity: a softer cluster can reorganize more easily toward nearby minima, amplifying differences associated with the functional, basis/ECP treatment, spin state, or numerical implementation [13,35].
For Sc4, this creates a dual scenario. Its structural adaptability and Lewis-acidic character may favor CO2 coordination, but the predicted adsorption behavior must be filtered carefully through vibrational validation and protocol comparison. Otherwise, the apparent accessibility of favorable adsorption geometries could reflect method-dependent stabilization rather than a robust structural feature.
2.12. Geometric Metrics for Sc4 D2h and Method and Spin Effect
The D2h-optimized Sc4 structures were analyzed through their Sc–Sc bond distances to quantify protocol-induced expansion and asymmetry relative to the initial geometry. These structural descriptors are summarized in Table 6.
Table 6.
Sc–Sc bond distances for the initial and optimized D2h Sc4 clusters calculated using different DFT protocols and spin multiplicities.
As reported in Table 6, the initial D2h Sc4 structure contains Sc–Sc distances in the range of 2.25–2.40 Å, whereas the optimized structures shift toward substantially longer metal–metal separations. In the singlet state, the optimized distances are mainly located around 2.79–2.81 Å, with moderate differences among the evaluated protocols. This systematic expansion indicates that the Sc4 framework is less compact than the corresponding Cu4 structures and that Sc–Sc bonding is more susceptible to relaxation during optimization.
The triplet structures exhibit greater asymmetry. Several Sc–Sc contacts approach or exceed 2.9 Å, particularly for r2SCAN-3c and TPSSh, indicating a more pronounced reorganization of the metal framework. This behavior suggests that spin-state changes affect the balance of Sc–Sc interactions and can reduce the cluster’s structural cohesion. Rather than preserving a rigid D2h arrangement, the triplet Sc4 structures tend to open or distort, generating a more flexible coordination environment.
This structural response is directly relevant for CO2 adsorption. An expanded Sc4 network can facilitate oxygen coordination and stabilize multidentate adsorption geometries, which is consistent with a polarization-dominated Lewis acid–base interaction mechanism. At the same time, the greater structural flexibility may increase Eads’s sensitivity to the computational protocol, because adsorption can be coupled to cluster reorganization. In this sense, Table 6 links the intrinsic geometry of Sc4 to the expected adsorption behavior and supports the need to interpret adsorption energies in conjunction with structural and vibrational validation [36,37,38].
2.13. Loss of Symmetry in Sc4 D3h as a Soft Surface Indicator
The initial and optimized D3h Sc4 geometries were compared to evaluate symmetry breaking and possible relaxation toward lower-symmetry structures, as summarized in Figure 8.
Figure 8.
Initial and optimized structures of Sc4 clusters with D3h starting symmetry, considering singlet and triplet multiplicities across the evaluated DFT protocols.
Figure 8 evidences that Sc4 does not consistently retain the D3h motif after structural relaxation. Several optimized geometries depart from the initial high-symmetry arrangement and evolve toward distorted or lower-symmetry structures. This behavior is consistent with a flexible metal framework and with the presence of nearby minima on a relatively shallow potential energy surface.
The loss of symmetry can be interpreted as a structural response to electron redistribution within the Sc4 cluster. Instead of preserving equivalent Sc–Sc interactions, the optimized structures tend to develop uneven metal–metal contacts, suggesting multicenter stabilization and partial opening of the framework. This behavior is characteristic of small early-transition-metal aggregates, where the balance between metal–metal cohesion, spin state, and geometric relaxation can be strongly method-dependent.
For CO2 adsorption, the structural plasticity observed in Figure 8 has two implications. On the one hand, a flexible Sc4 framework may accommodate diverse adsorption motifs and facilitate oxygen-centered or multidentate coordination. On the other side, this same flexibility increases the risk that adsorption trends depend on the optimized cluster basin selected by each computational protocol. For this reason, the D3h-derived Sc4 structures require careful vibrational validation before their adsorption energies are compared or interpreted as evidence of catalytic activation [13,35,39].
2.14. Geometric Metrics for Sc4 D3h and Asymmetry Quantification
The Sc–Sc distances of the D3h-derived Sc4 structures were analyzed to quantify the extent of symmetry breaking and structural rearrangement after optimization. The corresponding structural descriptors are summarized in Table 7.
Table 7.
Sc–Sc bond distances for the initial and optimized D3h Sc4 clusters calculated using different DFT protocols and spin multiplicities.
The data collected in Table 7 show that the initial D3h pattern is not preserved after optimization. Several protocols generate markedly uneven Sc–Sc distances, and in some cases, the longest contacts approach 3.0 Å. In contrast, other Sc–Sc separations remain in the approximate range of 2.63–2.85 Å, indicating that the optimized structures undergo nonuniform reorganization rather than simple isotropic expansion.
Within this set, the PBE0-SDD singlet structure exhibits one of the clearest distance separations, consistent with pronounced distortion of the original D3h motif. A similar trend is observed for the PBE-D4 triplet case, where multiple Sc–Sc contacts fall in the 2.85–3.00 Å range. The r2SCAN-3c and TPSSh results also display two distance scales, suggesting partially open frameworks and incomplete retention of the starting connectivity pattern.
From an adsorption perspective, these structural changes are relevant because they may generate more exposed or coordinatively unsaturated regions, which can favor Lewis-type interactions with CO2. At the same time, Table 7 indicates that some entries appear as hyphens, reflecting that the effective symmetry or connectivity of the optimized structure has changed. For this reason, the adsorption analysis should not rely only on metal–metal distances. A more reliable interpretation requires additional descriptors of the adsorbed CO2 geometry, particularly the O–C–O angle and the C–O bond lengths, together with the corresponding energetic and vibrational criteria.
2.15. Sc4 CO2 Geometries and Conceptual Comparison of Interaction Mechanism
The optimized Sc4–CO2 structures were examined across different adsorption modes to relate the electropositive character of scandium to the observed coordination patterns and possible CO2 activation pathways, as summarized in Figure 9.
Figure 9.
Optimized Sc4–CO2 adsorption geometries obtained from different initial symmetries and computational methods. The numbers below each structure identify the optimized systems generated from the evaluated initial symmetries (D2h, D3h, and Td), spin multiplicities, and four computational methods: r2SCAN-3C, PBE0-SDD, PBE-D4/def2-TZVP, and TPSSh/def2-TZVP. Structures 13–24 correspond to singlet systems and structures 37–48 to triplet systems. Atom colors: Sc, purple; C, yellow; O, red.
The set of structures collected in Figure 9 reveals a broader diversity of adsorption motifs for Sc4–CO2 than that observed for the Cu4 analogs. In these complexes, the interaction is expected to be governed primarily by the Lewis-acid character of scandium and by polarization of the adsorbate, rather than by strong metal-to-CO2 back-donation. This behavior is consistent with the preference for oxygen-centered coordination and with the appearance of geometries in which CO2 undergoes bending or selective C–O elongation.
Another relevant feature of Figure 9 is the Sc4 framework’s structural adaptability. Several optimized complexes show that the cluster can reorganize substantially to accommodate the adsorbate, thereby generating a wide variety of minima and coordination environments. Such flexibility may be advantageous for CO2 binding because it facilitates multidentate interactions and stabilizes activated configurations. At the same time, this same property makes the predicted adsorption mode more sensitive to the computational protocol. It increases the need for careful discrimination between genuine activation and method-dependent overbinding [13,15,39].
For that reason, Figure 9 should be interpreted together with Table 7 and Table 8. The structural motifs alone are insufficient to distinguish weak molecular adsorption from highly activated configurations. A reliable interpretation requires combining the optimized geometry with charge-transfer analysis and energetic data, especially in cases where very negative adsorption energies could reflect either strong Lewis-type stabilization or protocol-induced bias. From an applied perspective, Sc4 offers a useful model for identifying active-site motifs that can stabilize moderately activated CO2, which may be relevant to the design of lightweight or doped materials for capture and pre-activation in CCU-related systems.
Table 8.
NPA charge-transfer analysis for CO2 adsorption on Sc4 singlet complexes. qCO2 was obtained by summing the NPA charges of the carbon and oxygen atoms of the adsorbed CO2 molecule, whereas qSc4 corresponds to the net charge of the Sc4 metallic fragment. Negative qCO2 values indicate electron accumulation on CO2, while values close to zero suggest weak adsorption with limited electronic activation. Rows showing non-neutral fragment-charge balance due to ECP/NPA charge conventions were retained only as qualitative records and were not directly compared with all-electron/def2-based protocols.
The NPA results summarized in Table 8 indicate that the charge-transfer response of Sc4–CO2 singlet complexes depends strongly on the initial symmetry and the computational protocol. For the D2h structures, r2SCAN-3c and PBE-D4/def2-TZVP produce qCO2 values close to zero, indicating negligible net electron transfer from the Sc4 framework to the adsorbed CO2 molecule. This behavior is consistent with their weak or nearly thermoneutral Eads values and suggests that the D2h singlet arrangement does not provide clear electronic activation of CO2 under these protocols.
A different behavior is observed for the D3h singlet structures. In Table 8, r2SCAN-3c gives qCO2 = −1.076 e, whereas PBE-D4/def2-TZVP gives qCO2 = −1.632 e. These strongly negative values indicate substantial electron accumulation on CO2, supporting the formation of electronically activated adsorption states. This interpretation agrees with the more negative Eads values calculated for these configurations, particularly for PBE-D4/def2-TZVP. However, the simultaneous presence of very negative qCO2 and strongly exergonic Eads should be treated cautiously, because it may reflect genuine CO2 activation but may also indicate protocol-dependent overbinding or excessive stabilization of a highly reorganized adsorption geometry [33,34,40].
The TPSSh D3h singlet case provides an important counterexample. Although Table 8 indicates charge transfer toward CO2, the corresponding Eads value is positive. This result shows that charge accumulation on CO2 alone is not sufficient to define a thermodynamically favorable adsorption process. A reliable assignment of CO2 activation must combine qCO2 with Eads, vibrational stability, and geometric descriptors such as O–C–O bending and C–O bond elongation.
The available Td singlet result corresponds only to PBE0-SDD, whose NPA charges are not directly comparable because of the ECP/NPA charge convention. This row was therefore retained only as a qualitative record and was not used to establish quantitative charge-transfer trends. Completing the Td comparison with all-electron/def2-based protocols would be necessary to evaluate whether the charge-transfer behavior observed for D3h also extends to three-dimensional Sc4-derived adsorption geometries.
2.16. Gibbs Free Energy Analysis and Thermodynamic Consistency of CO2 Adsorption
The adsorption energies of CO2 on Cu4 and Sc4 were compared within a single energetic profile to assess how catalyst identity, initial symmetry, computational protocol, and spin multiplicity affect the predicted adsorption strength. The complete comparison is presented in Figure 10.
Figure 10.
CO2 adsorption energies, Eads, calculated for Cu4 and Sc4 clusters as a function of catalyst type, initial symmetry, DFT protocol, and spin multiplicity.
The energetic profile in Figure 10 confirms that Cu4 generally remains within a weak or marginal adsorption regime. This behavior is especially evident for the D2h- and D3h-derived structures, where both singlet and triplet states cluster near zero and may shift from slightly exergonic to slightly endergonic depending on the computational protocol. Such small energetic variations are consistent with weak adsorption or incipient chemisorption, where the final Eads value is sensitive to the balance between dispersion, exchange–correlation treatment, and structural relaxation.
A different trend is observed for Td-derived Cu4 structures. In this case, the singlet state tends to produce more negative adsorption energies than the corresponding planar arrangements, suggesting that the three-dimensional topology may provide a more favorable local coordination environment for CO2 stabilization. This could arise from stronger metal–adsorbate cooperation or from a more suitable orbital arrangement for adsorbate polarization. The triplet state, however, remains closer to the weak-adsorption regime in several protocols, indicating that spin multiplicity modifies the electronic availability of the Cu4 framework toward CO2 binding [22,25,33].
For Sc4, Figure 10 reveals a broader and more irregular energetic distribution. Several D3h- and Td-derived cases show markedly exergonic adsorption, but the magnitude and even the sign of Eads depend strongly on the method and spin state. The presence of outliers, including a sign inversion for one D3h singlet case, indicates that Sc4 adsorption is highly sensitive to the selected computational protocol and to the optimized structural basin reached during relaxation.
Chemically, this larger dispersion is consistent with the greater electropositivity and structural deformability of scandium. These features can enhance Lewis acid–base interactions and CO2 polarization, but they also make the computed adsorption energy more sensitive to the cluster’s deformation energy and to how each functional treats polarization, dispersion, and metal–adsorbate stabilization. For this reason, the most negative Sc4 adsorption energies in Figure 10 should not be interpreted automatically as evidence of superior CO2 capture or activation. Within a benchmarking framework, they require additional validation using vibrational stability, optimized CO2 geometry, and charge-transfer descriptors to rule out protocol-dependent overbinding or convergence toward a different structural regime [38,41,42].
Cu4 with weak adsorption suggests potential for more reversible capture and lower regeneration penalty, while Sc4 may offer more intense stabilization compatible with pre-activation, but only if the prediction is reproducible and filtered by vibrational stability. This point is also relevant from methodological sustainability, because early identification of sensitivity by method and spin reduces computational false positives and avoids rework in simulation campaigns.
Figure 11 provides a consistency check for the thermochemical dataset by comparing the absolute Gibbs free energies of the isolated clusters and the corresponding cluster–CO2 complexes for both spin multiplicities. In both panels, the catalyst–CO2 complexes exhibit more negative absolute Gibbs free energies than the isolated catalysts, as expected, because they include an additional CO2 fragment. This trend confirms the dataset’s internal construction within each computational protocol.
Figure 11.
Absolute Gibbs free energies of the isolated catalysts and catalyst–CO2 complexes calculated for Cu4 and Sc4 clusters as a function of initial symmetry, DFT protocol, and spin multiplicity. Panel (a) corresponds to multiplicity 1 and panel (b) to multiplicity 3.
The comparison between panels (a) and (b) also evidences the effect of spin multiplicity on the absolute Gibbs free-energy scale. Variations between singlet and triplet states reflect the spin–structure coupling characteristic of low-nuclearity metal clusters, where changes in electron occupancy can modify both relative stability and the degree of geometric reorganization. However, Figure 11 should not be interpreted as a direct comparison of stability across different DFT protocols. The large offsets in absolute G values are mainly determined by the electronic-structure model, basis set, and ECP treatment, particularly in PBE0-SDD, where the energy reference is not directly comparable with all-electron/def2-based calculations [13,19,35].
For this reason, the chemical interpretation must rely on internally derived quantities, such as Eads and ΔGads, where systematic contributions partially cancel within the same protocol. In this context, Figure 11 functions as a traceability control rather than as a standalone adsorption-stability descriptor. It supports the subsequent use of adsorption energies, Gibbs-corrected adsorption quantities, vibrational filters, and dispersion metrics as more appropriate criteria for evaluating CO2 binding and activation.
2.17. Adsorption Energies and Direct Comparison Between Cu4 and Sc4
To complement the electronic adsorption-energy profile and include a thermodynamic layer in the analysis, Table 9 reports the Gibbs free energies of the isolated catalysts, the corresponding catalyst–CO2 complexes, and the calculated CO2 adsorption energies. This comparison allows the adsorption trends to be evaluated while keeping the energy quantities internally consistent within each computational protocol.
Table 9.
Gibbs free energies of the isolated catalysts and catalyst–CO2 complexes, together with the corresponding CO2 adsorption energies calculated for Cu4 and Sc4 clusters using different initial symmetries, spin multiplicities, and DFT protocols.
The thermochemical data summarized in Table 9 provide an additional basis for interpreting CO2 adsorption beyond the structural and charge-transfer descriptors discussed above. The Gibbs free energies of the isolated catalyst, Gcat, and the catalyst–CO2 complex, Gcat + CO2, follow the expected internal trend within each computational protocol, with the complex showing a more negative total free energy due to the incorporation of CO2. However, these absolute Gibbs free-energy values should not be compared directly across different DFT protocols, because each functional/basis-set/ECP combination defines a different electronic-energy reference. The chemically meaningful comparison is therefore obtained from derived adsorption quantities, where systematic contributions partially cancel within the same protocol.
For Cu4, Table 9 confirms that most planar D2h and D3h cases remain within a weak adsorption regime. Several adsorption energies are close to thermoneutrality or only slightly exergonic/endergonic, indicating that the Cu4–CO2 interaction is generally weak and strongly dependent on the computational protocol [22,24,25]. This behavior is consistent with the structural and NPA analyses, where Cu4 does not systematically induce strong CO2 stabilization or charge-transfer-driven activation [33,34]. The T_d singlet configurations show more negative adsorption values than the planar analogs, suggesting that the three-dimensional arrangement can provide a more favorable coordination environment for CO2 binding [22,25].
For Sc4, the adsorption energies listed in Table 9 span a much broader range. Several D3h and Td configurations show markedly negative values, especially for selected PBE-D4/def2-TZVP and TPSSh/def2-TZVP cases. This trend is chemically consistent with the greater electropositivity and Lewis acidity of scandium, which can enhance CO2 polarization and stabilize oxygen-coordinated or multidentate adsorption motifs. The interpretation, however, must remain cautious. Highly negative adsorption energies may reflect stronger CO2 binding. Still, they may also include contributions from extensive cluster reorganization, changes in the final optimized geometry, or protocol-dependent stabilization of a particular adsorption basin.
The multiplicity comparison in Table 9 further shows that the spin state must be included in the adsorption model. In Cu4, triplet cases remain mostly close to weak adsorption, whereas singlet Td configurations become more exergonic. In Sc4, several triplet configurations preserve negative adsorption values across the evaluated symmetries, while the singlet cases show larger method-dependent variation, including both strongly favorable and unfavorable adsorption values. This reinforces that adsorption thermodynamics in low-nuclearity clusters cannot be separated from the spin-state landscape or from the structural relaxation pathway.
Taken together with the NPA results, Table 9 also shows why adsorption energy alone is insufficient to assign CO2 activation. In several Cu4 cases, Eads is favorable or near thermoneutrality while qCO2 remains close to zero, indicating weak adsorption or polarization-dominated stabilization rather than charge-transfer-driven activation. Conversely, selected Sc4 D3h cases combine strongly negative qCO2 with exergonic Eads, supporting stronger electronic activation. However, the largest values should still be examined carefully because they may also reflect protocol-dependent overbinding or excessive structural reorganization. Thus, the revised analysis distinguishes CO2 activation from possible overbinding artifacts by requiring consistency among qCO2, Eads, vibrational stability, and optimized adsorption geometry.
2.18. Global Benchmark Uncertainty at Eads and Practical Recommendation
The global dispersion of Eads was quantified to estimate the methodological variability associated with the use of different DFT protocols across the complete Cu4/Sc4 adsorption dataset. The resulting statistical descriptors are summarized in Table 10.
Table 10.
Global statistical dispersion of CO2 adsorption energy values, Eads, calculated for the complete Cu4/Sc4 dataset. The table reports the mean absolute deviation, standard deviation, and maximum absolute error in eV.
As summarized in Table 10, the CO2 adsorption-energy dataset exhibits substantial global dispersion across the evaluated computational protocols. The mean absolute deviation of 1.82 eV indicates that, on average, the calculated Eads values deviate considerably from the selected reference level. Likewise, the standard deviation of 2.21 eV confirms a broad spread in the predicted adsorption energies. In comparison, the maximum absolute error of 7.41 eV evidences the presence of extreme deviations within the dataset. These deviations may originate from differences in optimized geometry, spin-state stabilization, or convergence toward different stationary points on shallow potential-energy surfaces [43].
Because the values in Table 10 correspond to global statistics for the complete Cu4/Sc4 dataset, they should not be used to assign the dispersion specifically to Cu4 or Sc4, nor to separate the individual effects of initial symmetry or spin multiplicity. The appropriate conclusion is therefore methodological: when all Cu4–CO2 and Sc4–CO2 calculations are considered together, Eads is strongly dependent on the computational protocol. Consequently, adsorption energy should not be interpreted as a standalone descriptor of adsorption strength without complementary structural, vibrational, and charge-transfer validation.
The main value of Table 10 is diagnostic. It quantifies the global uncertainty associated with the choice of DFT protocol and supports the use of additional acceptance criteria before comparing adsorption trends. In this context, optimized-geometry consistency, positive ωmin values, Gibbs free-energy corrections, and NPA-derived qCO2 descriptors are required to reduce the risk of overinterpreting isolated adsorption-energy values. This approach provides a more cautious and reproducible basis for evaluating CO2 adsorption and activation in low-nuclearity metal clusters.
3. Materials and Methods
3.1. Benchmark Design and Case Set
A benchmark set was designed to evaluate the sensitivity of CO2 adsorption on small metal clusters to the computational protocol, spin state, initial geometry, and vibrational stability. Cu4 and Sc4 were selected as model systems because they represent electronically contrasting metal clusters and exhibit structural and spin-state sensitivity during geometry optimization. The benchmark included the isolated Cu4 and Sc4 clusters, as well as the corresponding Cu4–CO2 and Sc4–CO2 adsorption complexes [22,25,35].
For each metal cluster, representative low-nuclearity topologies were considered, including different initial symmetry arrangements and plausible adsorption orientations of CO2. The initial CO2 configurations were selected to sample distinct interaction regions of the cluster and to avoid biasing the optimization toward a single adsorption mode [33]. The calculations were performed without imposing symmetry constraints during structural relaxation, allowing the clusters and cluster–CO2 complexes to reorganize freely toward the nearest stationary point. Singlet and triplet multiplicities were explored to account for the possible competition between low-lying spin states in these small transition-metal aggregates. The complete matrix of systems, initial geometries, spin multiplicities, and identifiers is summarized in Table 1.
3.2. DFT Protocols Evaluated
Four density functional theory (DFT) protocols were evaluated: TPSSh, r2SCAN-3c, PBE-D4/def2-TZVP, and PBE0-SDD. In this work, the term “DFT protocol” refers not only to the exchange–correlation functional but also to the associated basis set, effective core potential (ECP), dispersion correction, and, when applicable, internal numerical corrections. This definition was adopted because the functional and basis-set treatments are inseparable components of the computational description of metal clusters and adsorption complexes [16,21,44,45].
The r2SCAN-3c protocol was included as a low-cost composite DFT method that incorporates internal corrections designed to improve the robustness of geometries and relative energies. PBE-D4/def2-TZVP was selected as a generalized gradient approximation protocol with explicit D4 dispersion correction and a triple-zeta basis set. TPSSh was included as a meta-hybrid functional commonly used in transition-metal chemistry. In contrast, PBE0-SDD was used to evaluate the influence of a hybrid functional combined with an ECP-based description of the metal centers. This set of protocols was chosen to compare different levels of exchange–correlation treatment, dispersion description, and basis/ECP representation in the prediction of metal–CO2 interactions [46].
3.3. Compute Packets and Numerical Configuration
The calculations were performed using ORCA and Gaussian to evaluate the consistency of selected results between two quantum-chemistry packages. The purpose of this comparison was not to benchmark the overall capabilities of both programs, but to assess whether selected stationary points obtained under comparable computational settings showed consistent geometries, vibrational character, and minimum/saddle-point assignments [47].
In ORCA, geometry optimizations and frequency calculations were carried out using convergence criteria suitable for transition-metal clusters and metal–adsorbate complexes. Fine integration grids were used for the evaluated meta-GGA, hybrid, and dispersion-corrected protocols. When required by the specific method, auxiliary-basis or resolution-of-identity approximations were applied in accordance with the standard implementation of the corresponding protocol. In Gaussian, analogous SCF convergence criteria, geometry optimization thresholds, and integration-grid settings were used whenever possible to minimize numerical differences between implementations [48,49]. The ORCA–Gaussian comparison was restricted to protocols and cases that could be defined in a comparable manner in both codes. For cases where an exact one-to-one correspondence was not possible, the comparison was limited to stationary-point character, final geometry, and vibrational stability.
3.4. Geometric Optimization and Multiplicity Exploration
For each system in the benchmark set, full geometry optimizations were performed in the gas phase without symmetry restrictions. Different initial geometries and spin multiplicities were explored to account for the presence of multiple low-lying minima, which is characteristic of small metal clusters. For each protocol, optimized structures were evaluated according to their electronic energy, final geometry, and vibrational stability.
The lowest-energy structure was not accepted automatically as the representative candidate unless it also corresponded to a physically meaningful stationary point. Structures showing nonphysical dissociation of CO2, collapse of the metallic framework, or reorganization toward a topology unrelated to the intended starting motif were documented and, when appropriate, reoptimized from alternative initial geometries. In this way, the analysis considered not only the energetic ordering but also the structural consistency and physical validity of the optimized configurations [19].
3.5. Vibrational Validation and Minimum Criterion Using ωmin
The nature of each optimized structure was evaluated by harmonic vibrational frequency analysis. The lowest harmonic frequency of each structure, ωmin, was used as an operational descriptor of stationary-point stability. Structures with ωmin > 0 were assigned as vibrational minima within the harmonic approximation, indicating that all calculated frequencies were real. In contrast, structures with ωmin < 0 were considered to contain at least one imaginary frequency and were therefore assigned as unstable stationary points or possible saddle points [50,51].
For cases with small-magnitude imaginary frequencies, additional checks were performed to distinguish between numerical artifacts and genuine instabilities on the potential energy surface. These checks included reoptimization with stricter SCF, geometry convergence, and integration-grid criteria when necessary. The use of ωmin was particularly relevant because low-nuclearity metal clusters often display soft vibrational modes and shallow potential energy surfaces [52]. Therefore, vibrational validation was used as a mandatory criterion before comparing adsorption energies. The results were summarized in a classification table and visualized by group in the corresponding figure.
3.6. Adsorption Energy and Thermodynamic Conventions
Adsorption energy was defined as:
where E(cluster–CO2), E(cluster), and E(CO2) correspond to the electronic energies of the optimized adsorption complex, isolated metal cluster, and isolated CO2 molecule, respectively. All energies used in a given Eads calculation were obtained using the same DFT protocol and, when applicable, the same numerical treatment. This ensured internal consistency between the isolated fragments and the adsorption complex [22,39].
Eads = E(cluster–CO2) − E(cluster) − E(CO2)
In addition to electronic adsorption energies, zero-point energy (ZPE) and thermal Gibbs free-energy corrections were calculated for the vibrationally stable minima. These corrections were obtained from harmonic vibrational frequency calculations performed at the same level of theory as the corresponding geometry optimization. The Gibbs free energy of adsorption at 298.15 K was calculated as:
where G(cluster–CO2), G(cluster), and G(CO2) correspond to the Gibbs free energies at 298.15 K of the optimized adsorption complex, isolated metal cluster, and isolated CO2 molecule, respectively. Only structures confirmed as vibrational minima were considered for the thermodynamic comparison between Eads and ΔGads [40].
ΔGads = G(cluster–CO2) − G(cluster) − G(CO2)
The basis set superposition error (BSSE) was also considered because it can artificially stabilize weakly bound cluster–adsorbate complexes when finite basis sets are used. In the r2SCAN-3c protocol, BSSE is not removed through an explicit counterpoise calculation, but is reduced by the geometrical counterpoise correction included in the composite scheme. In contrast, TPSSh, PBE-D4/def2-TZVP, and PBE0-SDD do not include an intrinsic BSSE correction; therefore, their adsorption energies were interpreted consistently within each protocol [15,19]. When necessary, representative counterpoise calculations can be used to estimate the residual BSSE contribution and evaluate its effect on adsorption-energy trends.
3.7. NPA Charge-Transfer Analysis
Natural population analysis (NPA) was performed for the selected Cu4–CO2 and Sc4–CO2 adsorption complexes to evaluate the extent of charge transfer between the metal cluster and the adsorbed CO2 molecule. The net charge on CO2 was calculated as qCO2 = qC + qO1 + qO2, where qC and qO correspond to the NPA charges of the carbon and oxygen atoms of the adsorbed molecule [34]. The net charge of the metallic fragment was calculated as the sum of the NPA charges of the four metal atoms. Negative qCO2 values were interpreted as electron accumulation on CO2, whereas values close to zero were interpreted as negligible net charge transfer. These descriptors were analyzed together with Eads and the optimized adsorption geometry to distinguish electronically activated CO2 from weakly adsorbed or possibly overbound configurations.
3.8. Validation with Cu2 as a Physical-Chemical Reference
The Cu2 dimer was used as an auxiliary validation system to support the selection of a reference protocol through experimentally comparable observables. This validation was based on the Cu–Cu equilibrium distance, the harmonic vibrational frequency associated with the Cu–Cu stretching mode, the ionization potential (IP), and the electron affinity (EA). These properties were selected because they probe complementary aspects of the electronic and structural description of copper clusters, including metal–metal bonding, vibrational curvature, and charge-sensitive behavior.
For each evaluated protocol, the neutral Cu2 dimer was optimized, and its vibrational frequency was calculated. The IP and EA were obtained using an adiabatic approach, in which the neutral, cationic, and anionic species were independently optimized for their corresponding charge and spin states. The IP was calculated as the energy difference between the optimized cation and neutral species. In contrast, the EA was calculated as the energy difference between the optimized neutral and anionic species. The calculated values were compared with the available experimental references summarized in Table 3. The protocol that showed the best overall compromise among Cu–Cu distance, vibrational frequency, IP, and EA was used as an internal reference to evaluate the dispersion of adsorption energies in the Cu4–CO2 and Sc4–CO2 systems [53,54].
3.9. ORCA vs. Gaussian Cross-Validation and Discrepancy Diagnosis
Selected cases were cross-validated between ORCA and Gaussian to assess the reproducibility of the optimized stationary points. The comparison focused on final geometry, final symmetry, the lowest vibrational frequency, and the character of the stationary point, distinguishing between true minima and structures with imaginary frequencies. This analysis was restricted to selected representative cases and was not intended as a general comparison of the two software packages.
When discrepancies were observed, they were interpreted as possible differences in numerical implementation, including integration grids, SCF convergence behavior, optimization algorithms, treatment of exact exchange, auxiliary-basis approximations, and initial SCF guesses. Cases in which one code produced a positive ωmin while the other produced ωmin < 0 were analyzed as examples of sensitivity in shallow regions of the potential energy surface. These differences indicate that nominally similar computational protocols may converge to different local regions of the potential energy surface or may yield different Hessian curvatures for low-frequency modes. A representative comparison is shown in Figure 3.
3.10. Statistical Analysis of Dispersion and Uncertainty in Eads
The uncertainty associated with the choice of computational protocol was quantified using dispersion metrics applied to Eads. Deviations were calculated relative to the selected reference protocol according to:
where Eads(method) is the adsorption energy obtained with a given protocol and Eads(reference) is the adsorption energy obtained with the selected reference protocol for the same system, geometry class, and spin state. From the ΔEads values, the mean absolute deviation (MAD), standard deviation (STD), and maximum absolute error, max |ΔEads|, were calculated. These metrics were evaluated globally and, when appropriate, separately for Cu4 and Sc4 to identify whether the protocol-dependent dispersion was system-specific [55,56].
ΔEads = Eads (method) − Eadsreference)
This statistical analysis was used to quantify the extent to which adsorption energies depend on the computational protocol. In this way, the comparison does not rely only on qualitative differences between optimized structures but also provides a numerical estimate of methodological uncertainty. The resulting dispersion metrics support the selection of more reliable computational protocols for subsequent studies of CO2 adsorption and activation in small metal clusters.
4. Conclusions
This study evaluated the sensitivity of CO2 adsorption on Cu4 and Sc4 clusters to the computational protocol, spin multiplicity, initial geometry, and vibrational stability. The results confirm that low-nuclearity metal clusters should not be treated as rigid adsorption sites, because structural relaxation may induce symmetry breaking, convergence toward nearby minima, or stationary points with imaginary frequencies. For this reason, ωmin was used as a necessary acceptance criterion before comparing adsorption energies.
The Cu4 and Sc4 systems exhibited distinct adsorption behaviors. Cu4 generally showed weak or moderate CO2 interactions, especially for D2h- and D3h-derived structures, whereas more favorable adsorption was observed for selected Td configurations. In contrast, Sc4 displayed broader energetic dispersion and stronger structural reorganization, reflecting the greater sensitivity of its adsorption behavior to cluster deformation, electrostatic polarization, spin state, and exchange–correlation treatment. These findings indicate that Eads alone is insufficient to assign CO2 activation in small clusters and must be interpreted together with structural, vibrational, thermodynamic, and charge-transfer descriptors.
The Cu2 reference calculations provided an auxiliary experimental anchor for evaluating the computational protocols using Cu–Cu bond distances, vibrational frequencies, ionization potentials, and electron affinities. However, this validation was used only as a physical–chemical reference and not as a direct substitute for tetramer-level experimental benchmarking. The ORCA–Gaussian comparison further showed that, even under comparable nominal settings, some optimized structures may differ in final symmetry or stationary-point character. This reinforces the need for explicit reporting of numerical settings, convergence behavior, and vibrational validation when studying soft potential-energy surfaces.
The NPA charge-transfer analysis introduced an electronic criterion to distinguish genuine CO2 activation from possible protocol-dependent overbinding. Several Cu4 cases showed favorable or near-thermoneutral adsorption energies with qCO2 values close to zero, indicating weak adsorption or polarization-dominated stabilization rather than charge-transfer-driven activation. Conversely, selected Sc4 configurations exhibited both negative qCO2 and exergonic adsorption energies, supporting stronger electronic activation. However, the most extreme values require cautious interpretation because they may also involve significant cluster reorganization or method-dependent stabilization.
Author Contributions
Conceptualization, K.O.-P., R.O.-T. and J.H.-F.; Methodology, K.O.-P., R.O.-T. and J.H.-F.; Software, K.O.-P., R.O.-T. and J.H.-F.; Validation, K.O.-P., R.O.-T. and J.H.-F.; Formal analysis, K.O.-P., R.O.-T. and J.H.-F.; Investigation, K.O.-P., R.O.-T. and J.H.-F.; Resources, K.O.-P., R.O.-T. and J.H.-F.; Data curation, K.O.-P., R.O.-T. and J.H.-F.; Writing—original draft, K.O.-P., R.O.-T. and J.H.-F.; Writing—review & editing, K.O.-P., R.O.-T. and J.H.-F.; Visualization, K.O.-P., R.O.-T. and J.H.-F.; Supervision, K.O.-P., R.O.-T. and J.H.-F.; Project administration, K.O.-P., R.O.-T. and J.H.-F.; Funding acquisition, K.O.-P., R.O.-T. and J.H.-F. All authors have read and agreed to the published version of the manuscript.
Funding
This research received no external funding.
Institutional Review Board Statement
Not applicable.
Informed Consent Statement
Not applicable.
Data Availability Statement
The original contributions presented in this study are included in the article. Further inquiries can be directed to the corresponding author.
Conflicts of Interest
The authors declare no conflict of interest.
References
- Podder, J.; Patra, B.R.; Pattnaik, F.; Nanda, S.; Dalai, A.K. a review of carbon capture and valorization technologies. Energies 2023, 16, 2589. [Google Scholar] [CrossRef] [Scilit]
- Alli, Y.A.; Oladoye, P.O.; Ejeromedoghene, O.; Bankole, O.M.; Alimi, O.A.; Omotola, E.O.; Olanrewaju, C.A.; Philippot, K.; Adeleye, A.S.; Ogunlaja, A.S. Nanomaterials as catalysts for CO2 transformation into value-added products: A review. Sci. Total. Environ. 2023, 868, 161547. [Google Scholar] [CrossRef] [Scilit]
- Jones, W.D. Carbon Capture and Conversion. J. Am. Chem. Soc. 2020, 142, 4955–4957. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Kätelhön, A.; Meys, R.; Deutz, S.; Suh, S.; Bardow, A. Climate change mitigation potential of carbon capture and utilization in the chemical industry. Proc. Natl. Acad. Sci. USA 2019, 116, 11187–11194. [Google Scholar] [CrossRef] [Scilit]
- Gabrielli, P.; Gazzani, M.; Mazzotti, M. The role of carbon capture and utilization, carbon capture and storage, and biomass to enable a Net-Zero-CO2 emissions chemical industry. Ind. Eng. Chem. Res. 2020, 59, 7033–7045. [Google Scholar] [CrossRef] [Scilit]
- Nagireddi, S.; Agarwal, J.R.; Vedapuri, D. Carbon dioxide capture, utilization, and sequestration: Current status, challenges, and future prospects for global decarbonization. ACS Eng. Au 2023, 4, 22–48. [Google Scholar] [CrossRef] [Scilit]
- Gautam, P.; Salunke, D.; Lad, D.; Gautam, A. Convergent synergy of carbon capture within the circular economy paradigm: A nexus for realizing multifaceted sustainable development goals. Carbon Res. 2025, 4, 3. [Google Scholar] [CrossRef] [Scilit]
- Wang, J.; Zhang, Q.; Zhang, Z.; He, S.; Liang, N.; Gao, Y.; Deng, K.; Wang, Y.; Zhou, J.; Wu, K. Atomic-Scale Insights into CO2 and H2O Co-Adsorption on Sr2Fe1.5Mo0.5O6 Surfaces: Role of Electronic Structure and Dual-Site Interactions. Catalysts 2025, 15, 884. [Google Scholar] [CrossRef] [Scilit]
- Santiago-Cuevas, A.-J.; Palacios-Cabrera, C.-B.; Tecuapa-Flores, E.D.; Bazany-Rodríguez, I.J.; Narayanan, J.; Padilla-Martínez, I.I.; Aguilar, C.A.; Thangarasu, P. CO2 adsorption by Carbon Quantum Dots/Metal ferrites (M = CO2+, NI2+, and ZN2+): Electrochemical and theoretical studies. ACS Omega 2025, 10, 13977–14000. [Google Scholar] [CrossRef] [Scilit]
- Dastider, S.G.; Haldar, K.K.; Banerjee, A.; Mondal, K. Role of Back-Donation Deciphering CO2 Adsorption Modes on 3 d -Transition Metals. ChemPhysChem 2025, 27, e202500538. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Asgari, M.; Jawahery, S.; Bloch, E.D.; Hudson, M.R.; Flacau, R.; Vlaisavljevich, B.; Long, J.R.; Brown, C.M.; Queen, W.L. An experimental and computational study of CO2adsorption in the sodalite-type M-BTT (M = Cr, Mn, Fe, Cu) metal–organic frameworks featuring open metal sites. Chem. Sci. 2018, 9, 4579–4588. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Halder, A.; Curtiss, L.A.; Fortunelli, A.; Vajda, S. Perspective: Size selected clusters for catalysis and electrochemistry. J. Chem. Phys. 2018, 148, 110901. [Google Scholar] [CrossRef] [Scilit]
- de Lara-Castells, M.P. An Ab Initio Journey toward the Molecular-Level Understanding and Predictability of Subnanometric Metal Clusters. Small Struct. 2024, 5, 2400147. [Google Scholar] [CrossRef] [Scilit]
- Mitrushchenkov, A.O.; de Lara-Castells, M.P. High-level ab initio evidence of bipyramidal Cu5 clusters as fluxional Jahn-Teller molecules. ChemPhysChem 2023, 24, e202300317. [Google Scholar] [CrossRef] [Scilit]
- Rehak, F.R.; Piccini, G.; Alessio, M.; Sauer, J. Including dispersion in density functional theory for adsorption on flat oxide surfaces, in metal–organic frameworks and in acidic zeolites. Phys. Chem. Chem. Phys. 2020, 22, 7577–7585. [Google Scholar] [CrossRef] [Scilit]
- Butera, V. Density functional theory methods applied to homogeneous and heterogeneous catalysis: A short review and a practical user guide. Phys. Chem. Chem. Phys. 2024, 26, 7950–7970. [Google Scholar] [CrossRef] [Scilit]
- Liao, X.; Lu, R.; Xia, L.; Liu, Q.; Wang, H.; Zhao, K.; Wang, Z.; Zhao, Y. Density Functional Theory for Electrocatalysis. Energy Environ. Mater. 2021, 5, 157–185. [Google Scholar] [CrossRef] [Scilit]
- Choudhary, K.; Wines, D.; Li, K.; Garrity, K.F.; Gupta, V.; Romero, A.H.; Krogel, J.T.; Saritas, K.; Fuhr, A.; Ganesh, P.; et al. JARVIS-Leaderboard: A large scale benchmark of materials design methods. npj Comput. Mater. 2024, 10, 93. [Google Scholar] [CrossRef] [Scilit]
- Bursch, M.; Mewes, J.; Hansen, A.; Grimme, S. Best-Practice DFT Protocols for Basic Molecular Computational Chemistry. Angew. Chem. Int. Ed. 2022, 61, e202205735. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Shi, B.X.; Rosen, A.S.; Schäfer, T.; Grüneis, A.; Kapil, V.; Zen, A.; Michaelides, A. An accurate and efficient framework for modelling the surface chemistry of ionic materials. Nat. Chem. 2025, 17, 1688–1695. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Cramer, C.J.; Truhlar, D.G. Density functional theory for transition metals and transition metal chemistry. Phys. Chem. Chem. Phys. 2009, 11, 10757–10816. [Google Scholar] [CrossRef] [Scilit]
- Gálvez-González, L.E.; Juárez-Sánchez, J.O.; Pacheco-Contreras, R.; Garzón, I.L.; Paz-Borbón, L.O.; Posada-Amarillas, A. CO2 adsorption on gas-phase Cu4−xPtx (x = 0–4) clusters: A DFT study. Phys. Chem. Chem. Phys. 2018, 20, 17071–17080. [Google Scholar] [CrossRef] [Scilit]
- Jug, K.; Zimmermann, B.; Calaminici, P.; Köster, A.M. Structure and stability of small copper clusters. J. Chem. Phys. 2002, 116, 4497–4507. [Google Scholar] [CrossRef] [Scilit]
- Jaque, P.; Toro-Labbé, A. Characterization of copper clusters through the use of density functional theory reactivity descriptors. J. Chem. Phys. 2002, 117, 3208–3218. [Google Scholar] [CrossRef] [Scilit]
- Ha, N.T.T.; Hue, V.T.M.; Trinh, B.C.; Ha, N.N.; Cam, L.M. Study on the Adsorption and Activation Behaviours of Carbon Dioxide over Copper Cluster (Cu4) and Alumina-Supported Copper Catalyst (Cu4/Al2O3) by means of Density Functional Theory. J. Chem. 2019, 2019, 4341056. [Google Scholar] [CrossRef] [Scilit]
- Alcalá-Varilla, L.A.; Ponnefz-Durango, R.E.; Seriani, N.; Araujo-Lopez, E.; Montoya, J.A. A DFT + U study on the stability of small CUN clusters (N = 3–6 atoms): Calculation of phonon frequencies. Condens. Matter 2023, 8, 81. [Google Scholar] [CrossRef] [Scilit]
- Schlegel, H.B. Optimization of equilibrium geometries and transition structures. J. Comput. Chem. 1982, 3, 214–218. [Google Scholar] [CrossRef] [Scilit]
- Peng, C.; Schlegel, H.B. Combining synchronous transit and Quasi-Newton methods to find transition states. Isr. J. Chem. 1993, 33, 449–454. [Google Scholar] [CrossRef] [Scilit]
- Li, C.-G.; Shen, Z.-G.; Hu, Y.-F.; Tang, Y.-N.; Chen, W.-G.; Ren, B.-Z. Insights into the structures and electronic properties of Cun+1 μ and CunS μ (n = 1–12; μ = 0, ±1) clusters. Sci. Rep. 2017, 7, 1345. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Wang, J. Structural, electronic, and magnetic properties ofScn(n=2–18)clusters from density functional calculations. Phys. Rev. B 2007, 75, 155422. [Google Scholar] [CrossRef] [Scilit]
- National Institute of Standards and Technology. CCCBDB Listing of Experimental Data Page 2. Computational Chemistry Comparison and Benchmark Database (CCCBDB), Standard Reference Database 101, Release 22, May 2022. Available online: https://cccbdb.nist.gov/exp2x.asp?casno=12190704 (accessed on 24 December 2025).
- Szalay, M.; Buzsáki, D.; Barabás, J.; Faragó, E.; Janssens, E.; Nyulászi, L.; Höltzl, T. Screening of transition metal doped copper clusters for CO2 activation. Phys. Chem. Chem. Phys. 2021, 23, 21738–21747. [Google Scholar] [CrossRef] [Scilit]
- Collacique, M.N.; Ocampo-Restrepo, V.K.; Da Silva, J.L.F. Ab initio investigation of the role of the d-states on the adsorption and activation properties of CO2 on 3d, 4d, and 5d transition-metal clusters. J. Chem. Phys. 2022, 156, 124106. [Google Scholar] [CrossRef] [Scilit]
- Gautam, S.; Dharamvir, K.; Goel, N. CO2 adsorption and activation over medium sized Cun (n=7, 13 and 19) clusters: A density functional study. Comput. Theor. Chem. 2013, 1009, 8–16. [Google Scholar] [CrossRef] [Scilit]
- Yuan, H.K.; Chen, H.; Ahmed, A.S.; Zhang, J.F. Density-functional study of Scn(n=2–16)clusters: Lowest-energy structures, electronic structure, and magnetism. Phys. Rev. B 2006, 74, 144434. [Google Scholar] [CrossRef] [Scilit]
- Li, X.; Chen, Y.; Basnet, P.; Luo, J.; Wang, H. Probing the properties of size dependence and correlation for tantalum clusters: Geometry, stability, vibrational spectra, magnetism, and electronic structure. RSC Adv. 2019, 9, 1015–1028. [Google Scholar] [CrossRef] [Scilit]
- Sajjad, S.; Maria; Mahmood, T.; Ayub, K. Benchmark study of structural and vibrational properties of scandium clusters. J. Mol. Struct. 2017, 1142, 139–147. [Google Scholar] [CrossRef] [Scilit]
- Pangh, A. Catalytic cleavage of CO2 on bimetallic Ni4M (M = Ni, Mo, Sc, and Y) nanoclusters: A DFT study. Catal. Commun. 2020, 149, 106245. [Google Scholar] [CrossRef] [Scilit]
- Montejo-Alvaro, F.; Martínez-Espinosa, J.A.; Rojas-Chávez, H.; Navarro-Ibarra, D.C.; Cruz-Martínez, H.; Medina, D.I. CO2 Adsorption over 3d Transition-Metal Nanoclusters Supported on Pyridinic N3-Doped Graphene: A DFT Investigation. Materials 2022, 15, 6136. [Google Scholar] [CrossRef] [Scilit]
- Sillar, K.; Koppel, I. AB Initio Calculation of thermodynamic functions for CO2 adsorption in Metal–Organic frameworks: Entropic effects of lateral interactions. J. Phys. Chem. C 2023, 127, 11712–11719. [Google Scholar] [CrossRef] [Scilit]
- Liu, P.; Han, J.; Chen, Y.; Yu, H.; Zhou, X.; Zhang, W. Binding strengths and orientations in CO2 adsorption on cationic scandium oxides: Governing factor revealed by a combined infrared spectroscopy and theoretical study. J. Phys. Chem. A 2024, 128, 3007–3014. [Google Scholar] [CrossRef] [Scilit]
- Wu, G.; Wang, J.; Lu, Y.; Yang, M. Density functional study of CO adsorption on Scn (n=2–13) clusters. J. Chem. Phys. 2008, 128, 224315. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Araujo, R.B.; Rodrigues, G.L.S.; dos Santos, E.C.; Pettersson, L.G.M. Adsorption energies on transition metal surfaces: Towards an accurate and balanced description. Nat. Commun. 2022, 13, 6853. [Google Scholar] [CrossRef] [Scilit]
- Grimme, S.; Hansen, A.; Ehlert, S.; Mewes, J.-M. r2SCAN-3c: A ‘Swiss army knife’ composite electronic-structure method. J. Chem. Phys. 2021, 154, 064103. [Google Scholar] [CrossRef] [Scilit]
- Caldeweyher, E.; Ehlert, S.; Hansen, A.; Neugebauer, H.; Spicher, S.; Bannwarth, C.; Grimme, S. A generally applicable atomic-charge dependent London dispersion correction. J. Chem. Phys. 2019, 150, 154122. [Google Scholar] [CrossRef] [Scilit]
- Weymuth, T.; Couzijn, E.P.A.; Chen, P.; Reiher, M. New benchmark Set of Transition-Metal Coordination Reactions for the assessment of density functionals. J. Chem. Theory Comput. 2014, 10, 3092–3103. [Google Scholar] [CrossRef] [Scilit]
- Smith, D.G.A.; Lolinco, A.T.; Glick, Z.L.; Lee, J.; Alenaizan, A.; Barnes, T.A.; Borca, C.H.; Di Remigio, R.; Dotson, D.L.; Ehlert, S.; et al. Quantum Chemistry Common Driver and Databases (QCDB) and Quantum Chemistry Engine (QCEngine): Automation and interoperability among computational chemistry programs. J. Chem. Phys. 2021, 155, 204801. [Google Scholar] [CrossRef] [Scilit]
- Neese, F.; Wennmohs, F.; Becker, U.; Riplinger, C. The ORCA quantum chemistry program package. J. Chem. Phys. 2020, 152, 224108. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Weigend, F. Accurate Coulomb-fitting basis sets for H to Rn. Phys. Chem. Chem. Phys. 2006, 8, 1057. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Schlegel, H.B. Geometry optimization. Wiley Interdiscip. Rev. Comput. Mol. Sci. 2011, 1, 790–809. [Google Scholar] [CrossRef] [Scilit]
- Baker, J. An algorithm for the location of transition states. J. Comput. Chem. 1986, 7, 385–395. [Google Scholar] [CrossRef] [Scilit]
- Fielicke, A. Probing the binding and activation of small molecules by gas-phase transition metal clustersviaIR spectroscopy. Chem. Soc. Rev. 2023, 52, 3778–3841. [Google Scholar] [CrossRef] [Scilit]
- Balbuena, P.B.; Derosa, P.A.; Seminario, J.M. Density Functional Theory Study of Copper Clusters. J. Phys. Chem. B 1999, 103, 2830–2840. [Google Scholar] [CrossRef] [Scilit]
- Ho, J.; Ervin, K.M.; Lineberger, W.C. Photoelectron spectroscopy of metal cluster anions: Cu−n, Ag−n, and Au−n. J. Chem. Phys. 1990, 93, 6987–7002. [Google Scholar] [CrossRef] [Scilit]
- Goerigk, L.; Hansen, A.; Bauer, C.; Ehrlich, S.; Najibi, A.; Grimme, S. A look at the density functional theory zoo with the advanced GMTKN55 database for general main group thermochemistry, kinetics and noncovalent interactions. Phys. Chem. Chem. Phys. 2017, 19, 32184–32215. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Wellendorff, J.; Silbaugh, T.L.; Garcia-Pintos, D.; Nørskov, J.K.; Bligaard, T.; Studt, F.; Campbell, C.T. A benchmark database for adsorption bond energies to transition metal surfaces and comparison to selected DFT functionals. Surf. Sci. 2015, 640, 36–44. [Google Scholar] [CrossRef] [Scilit]
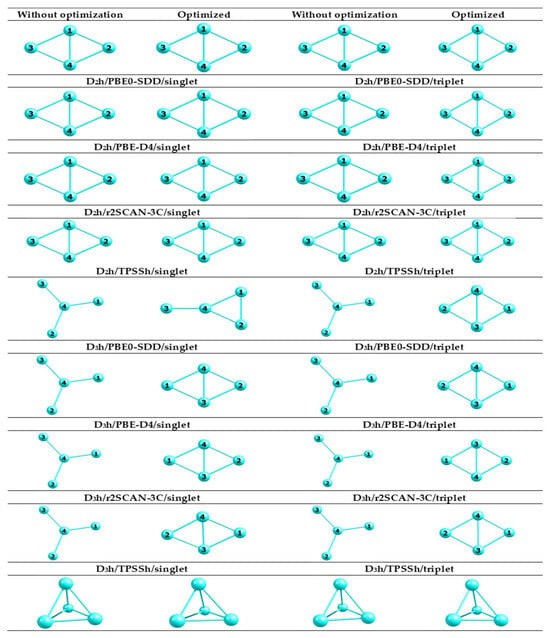
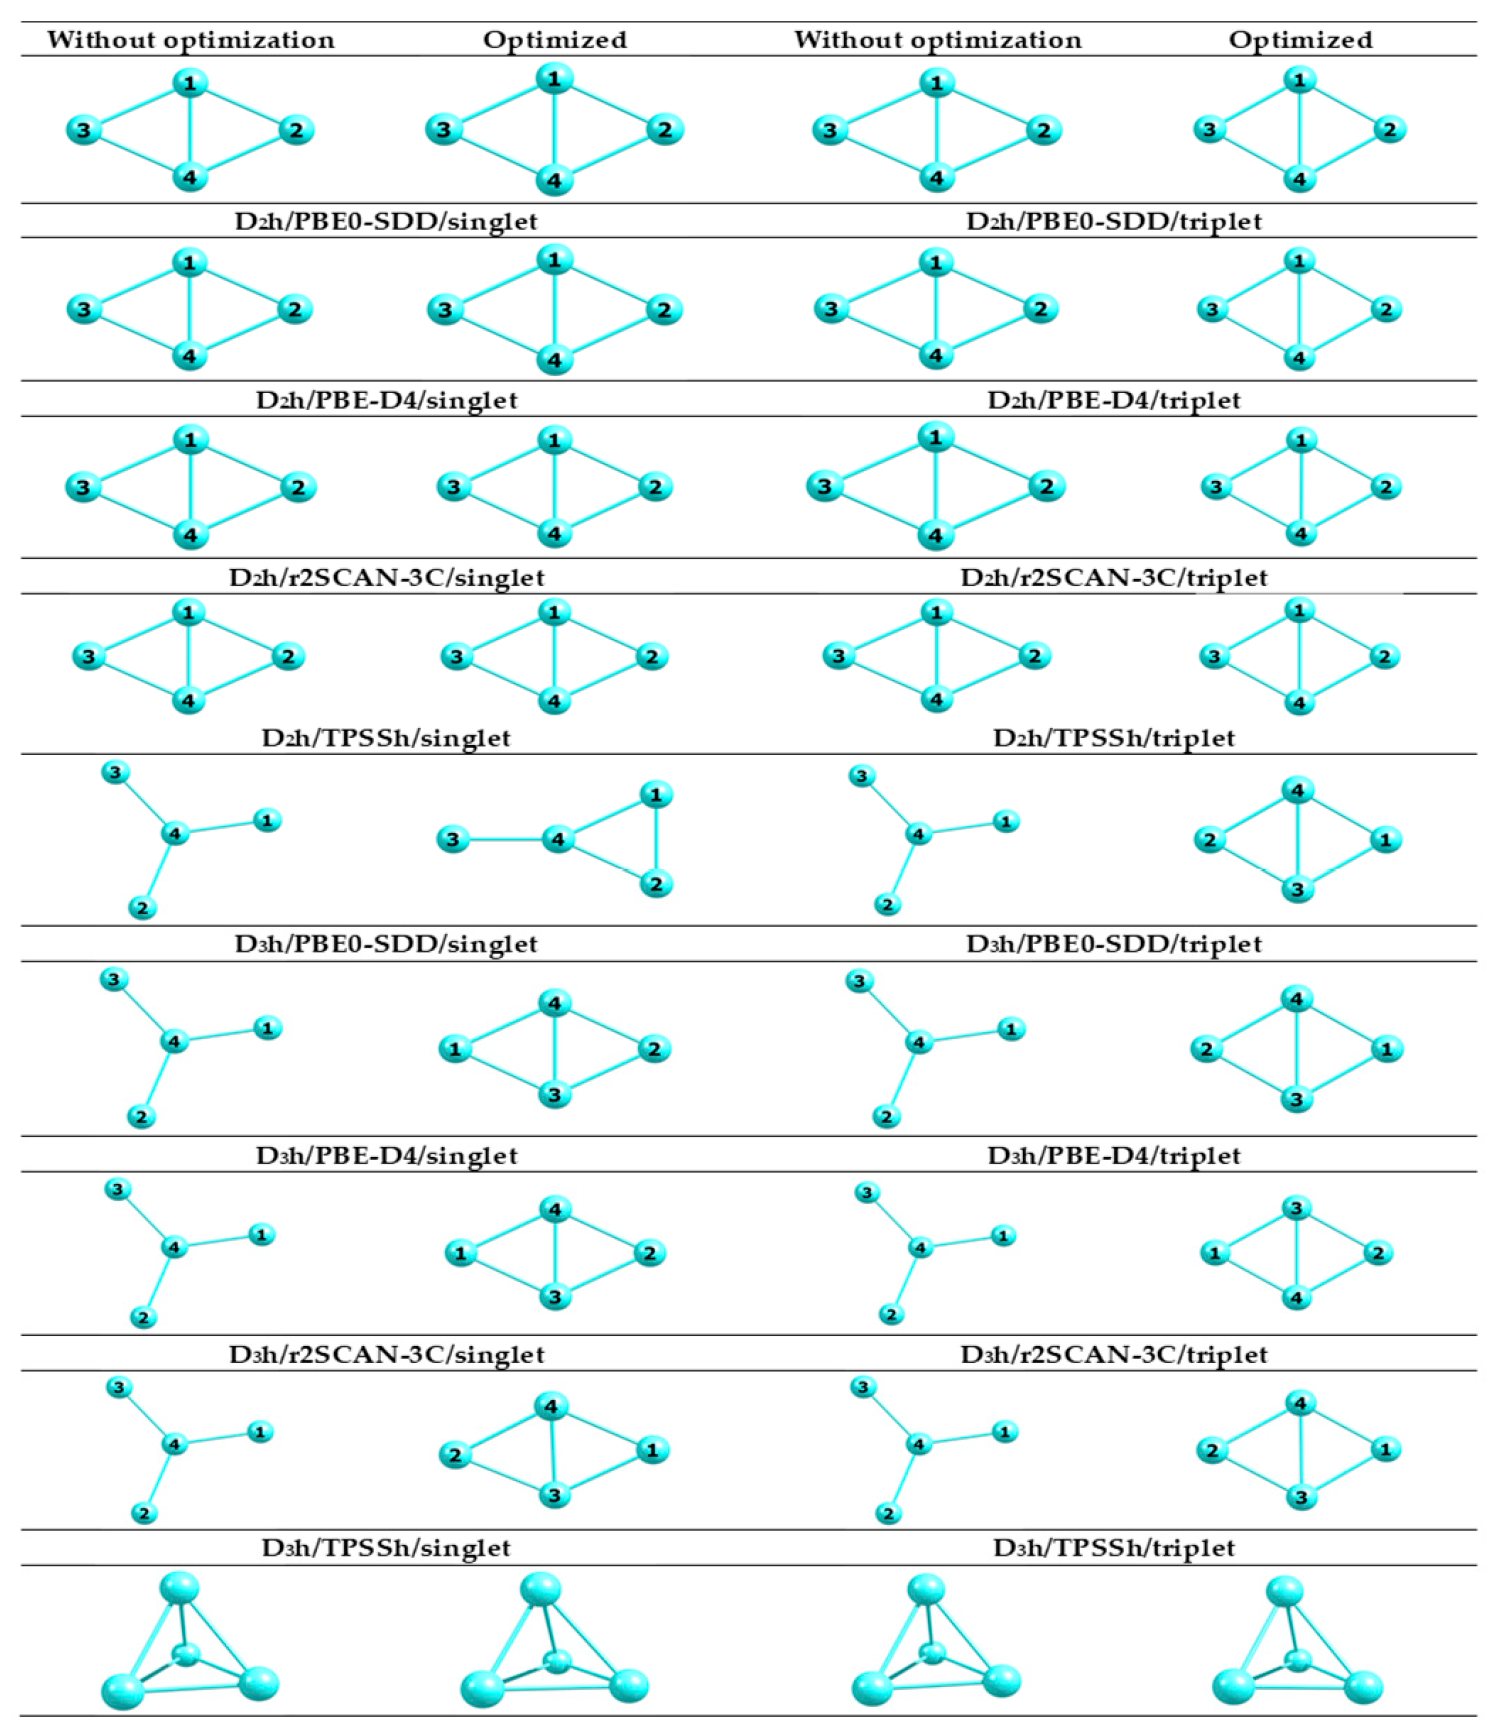
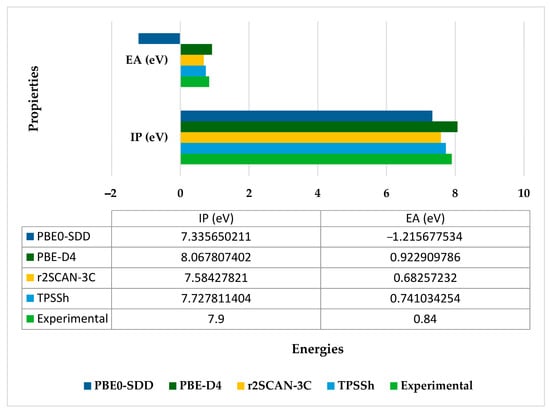
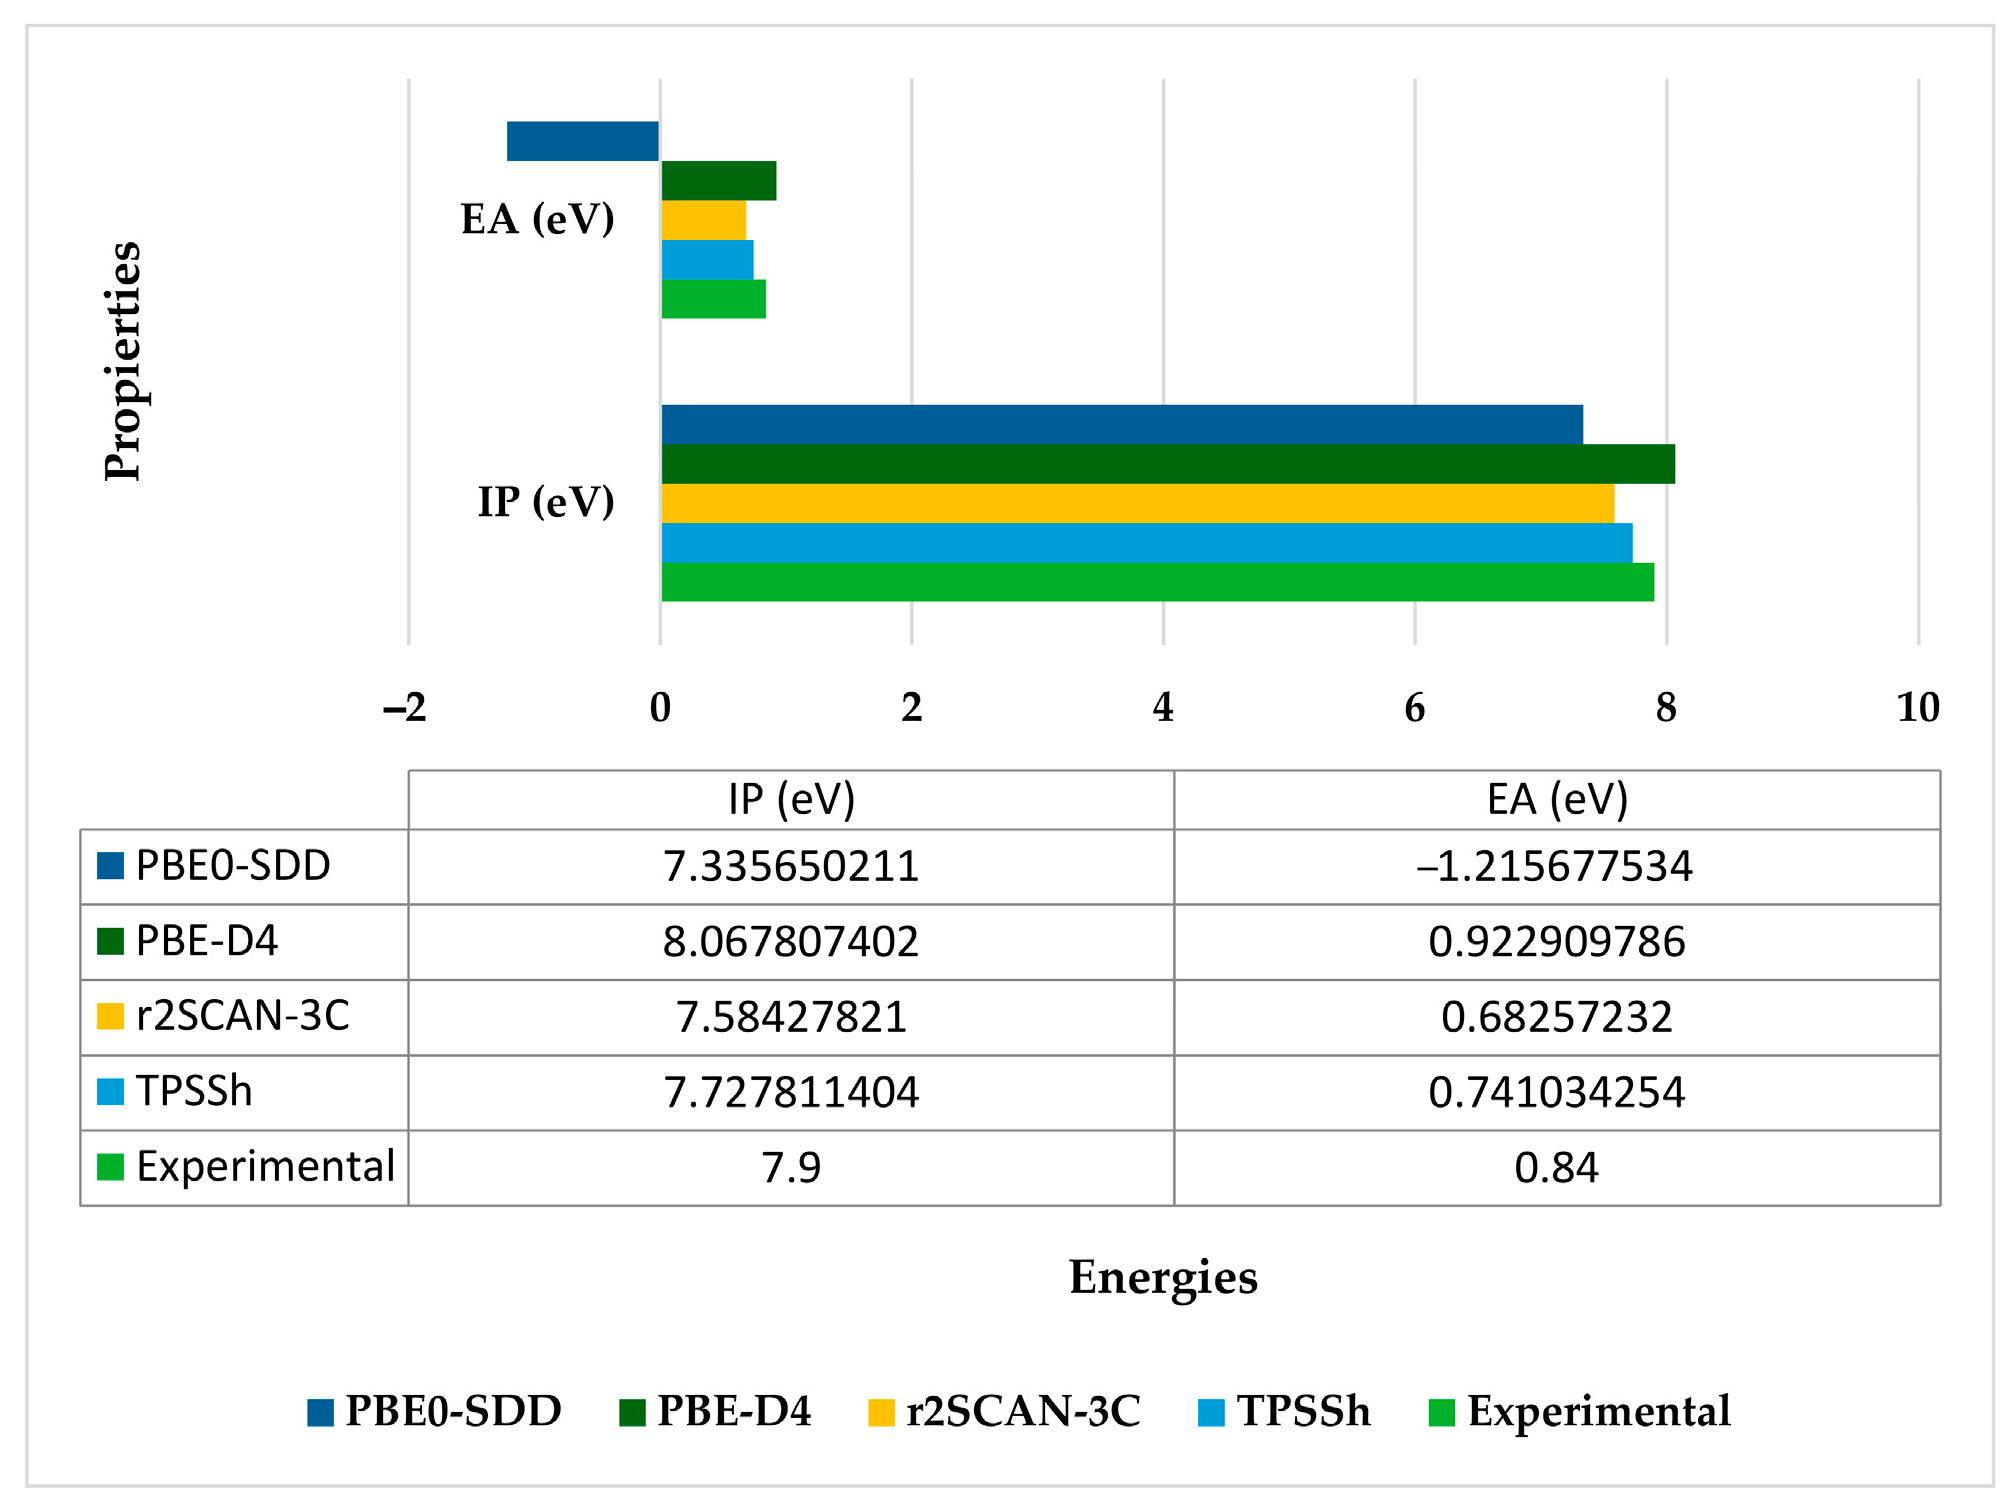
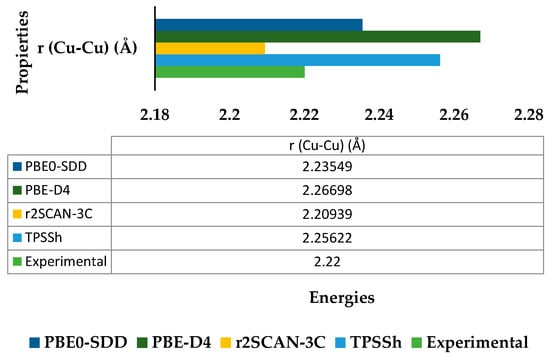
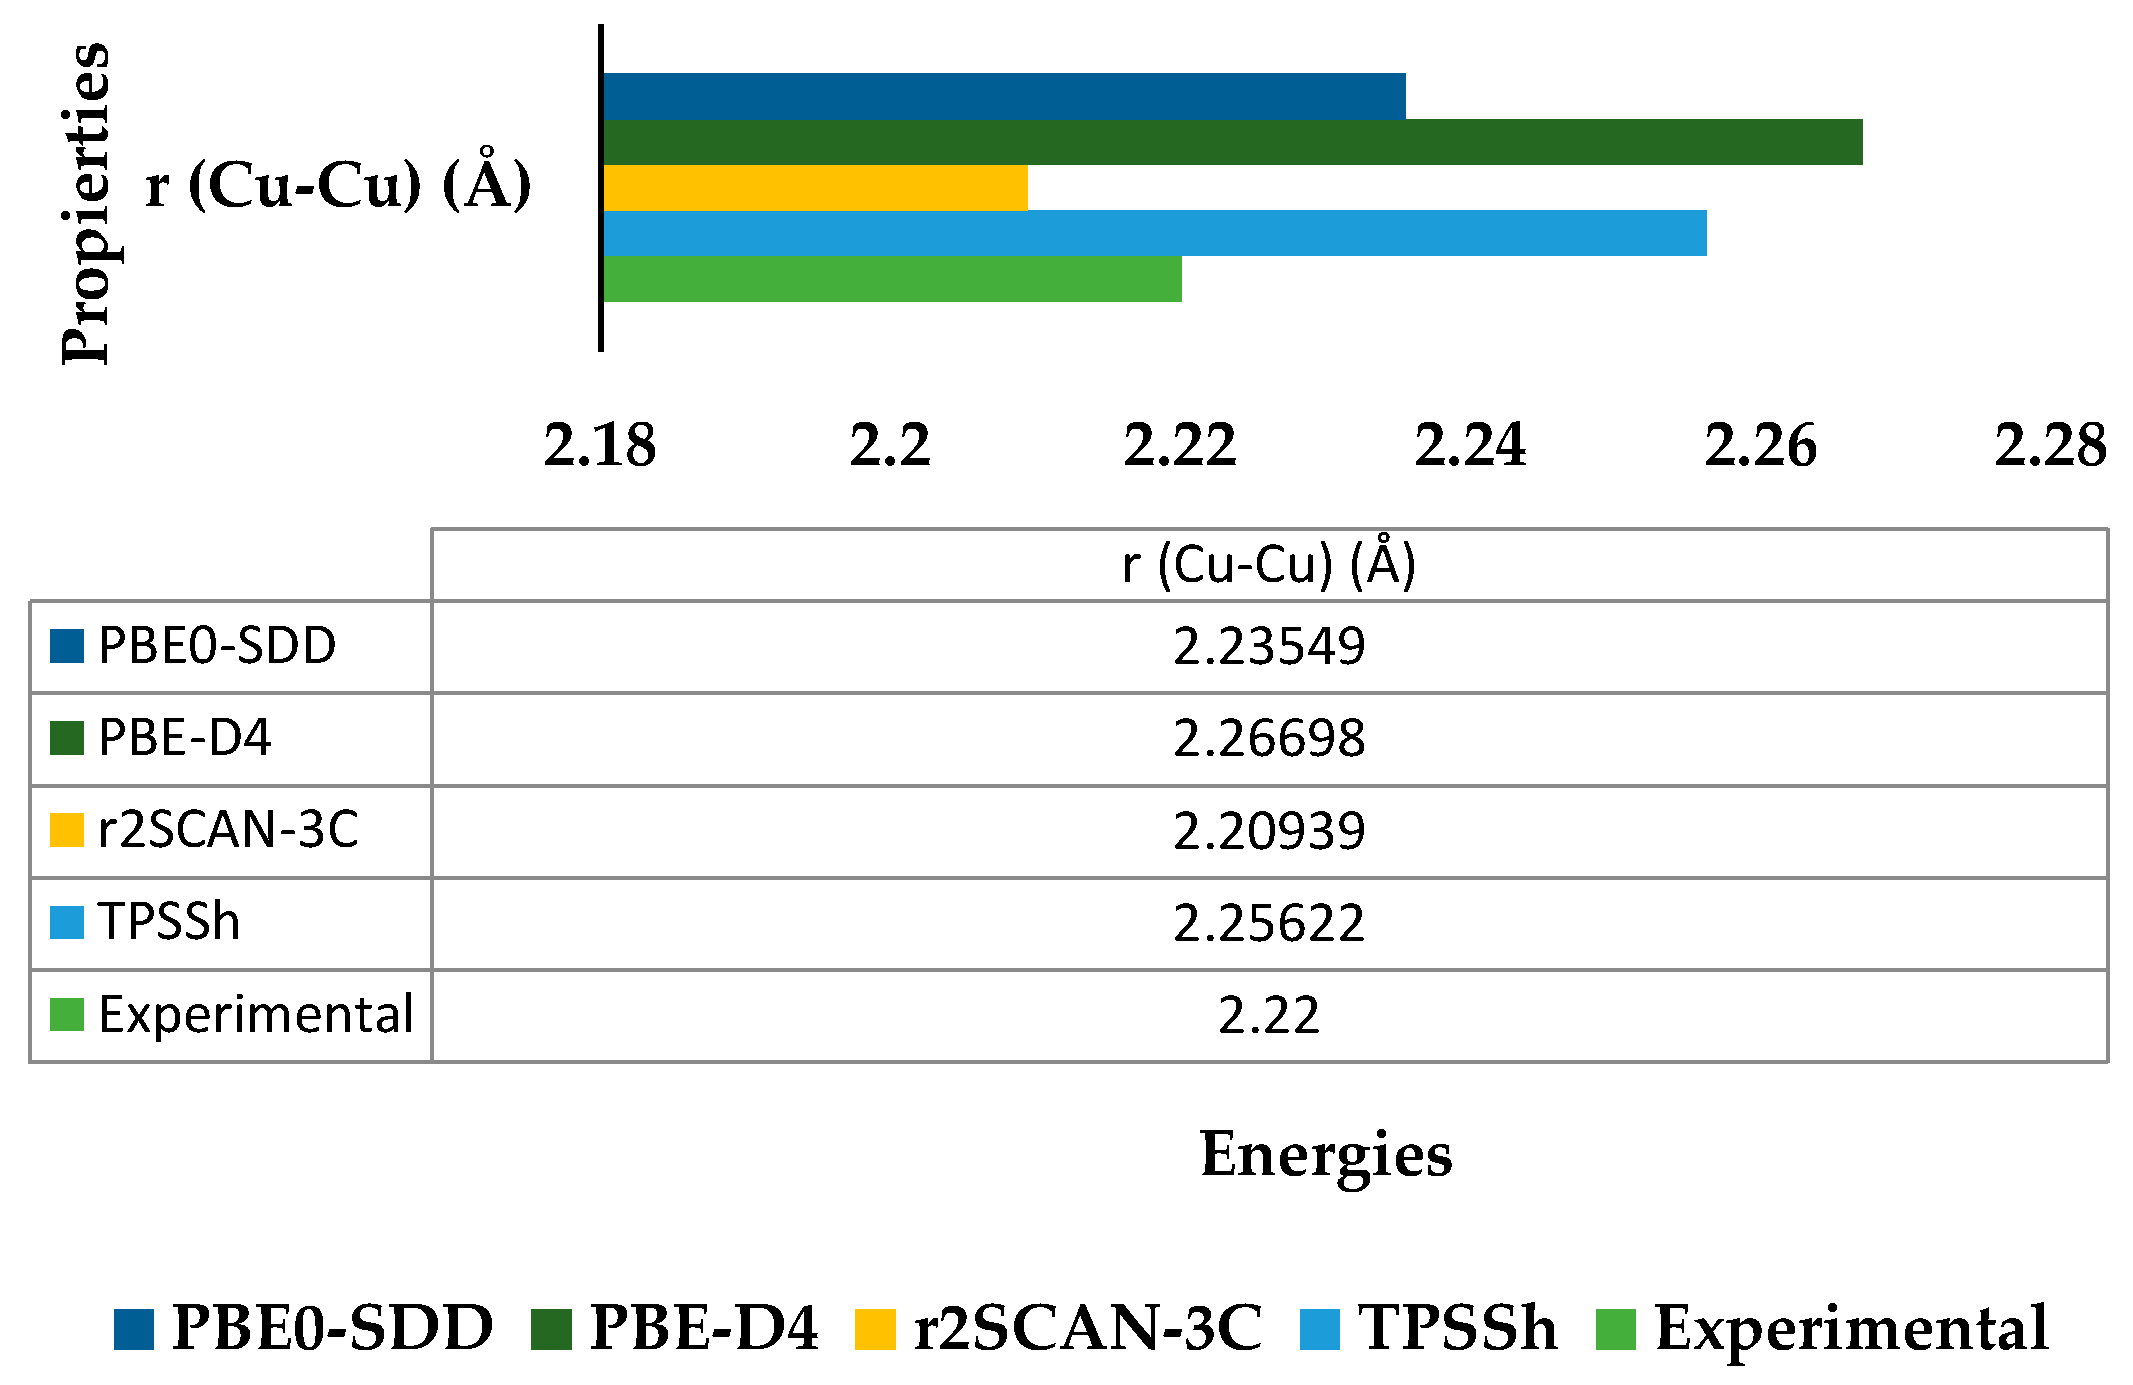
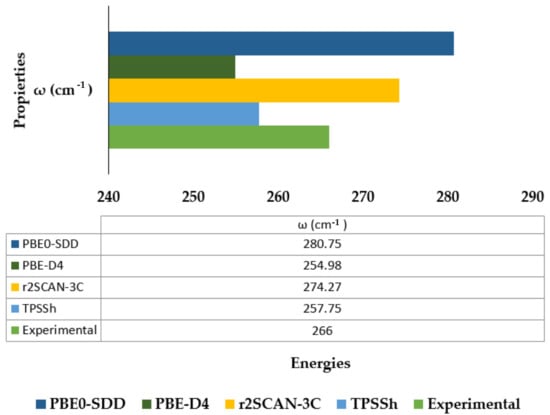
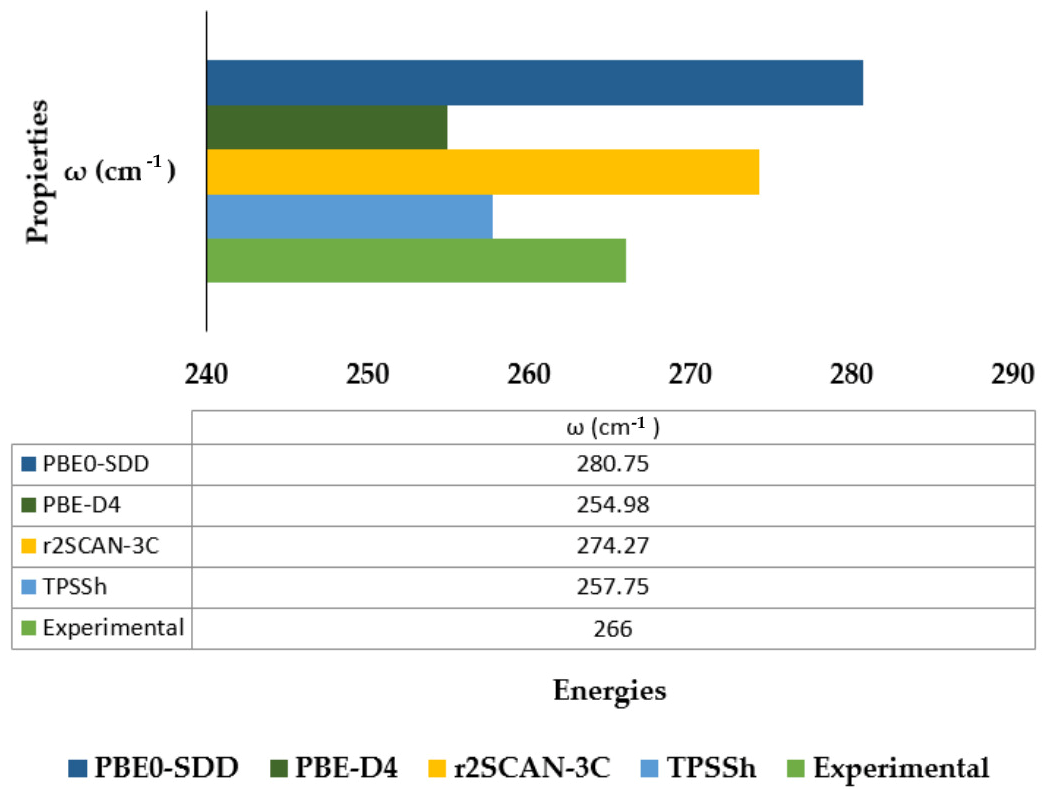
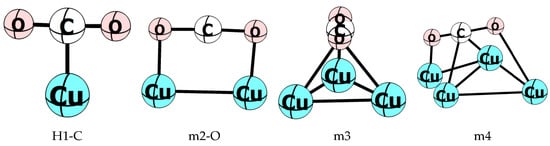
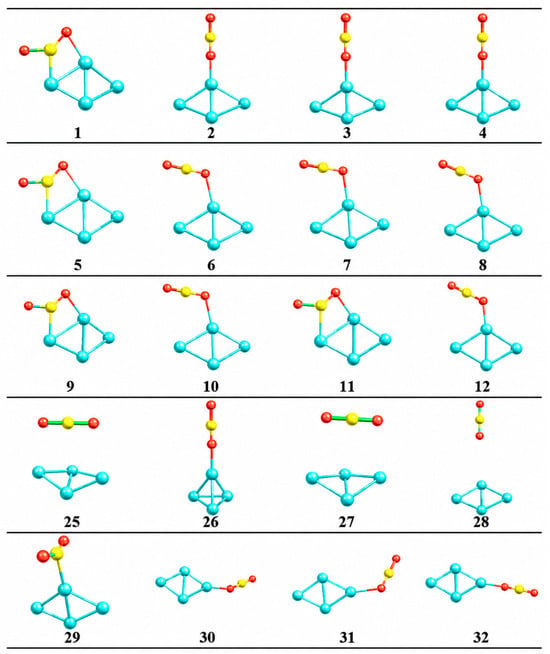
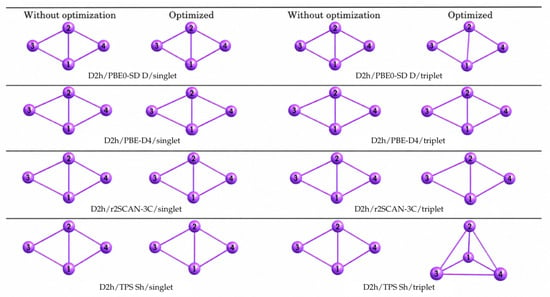
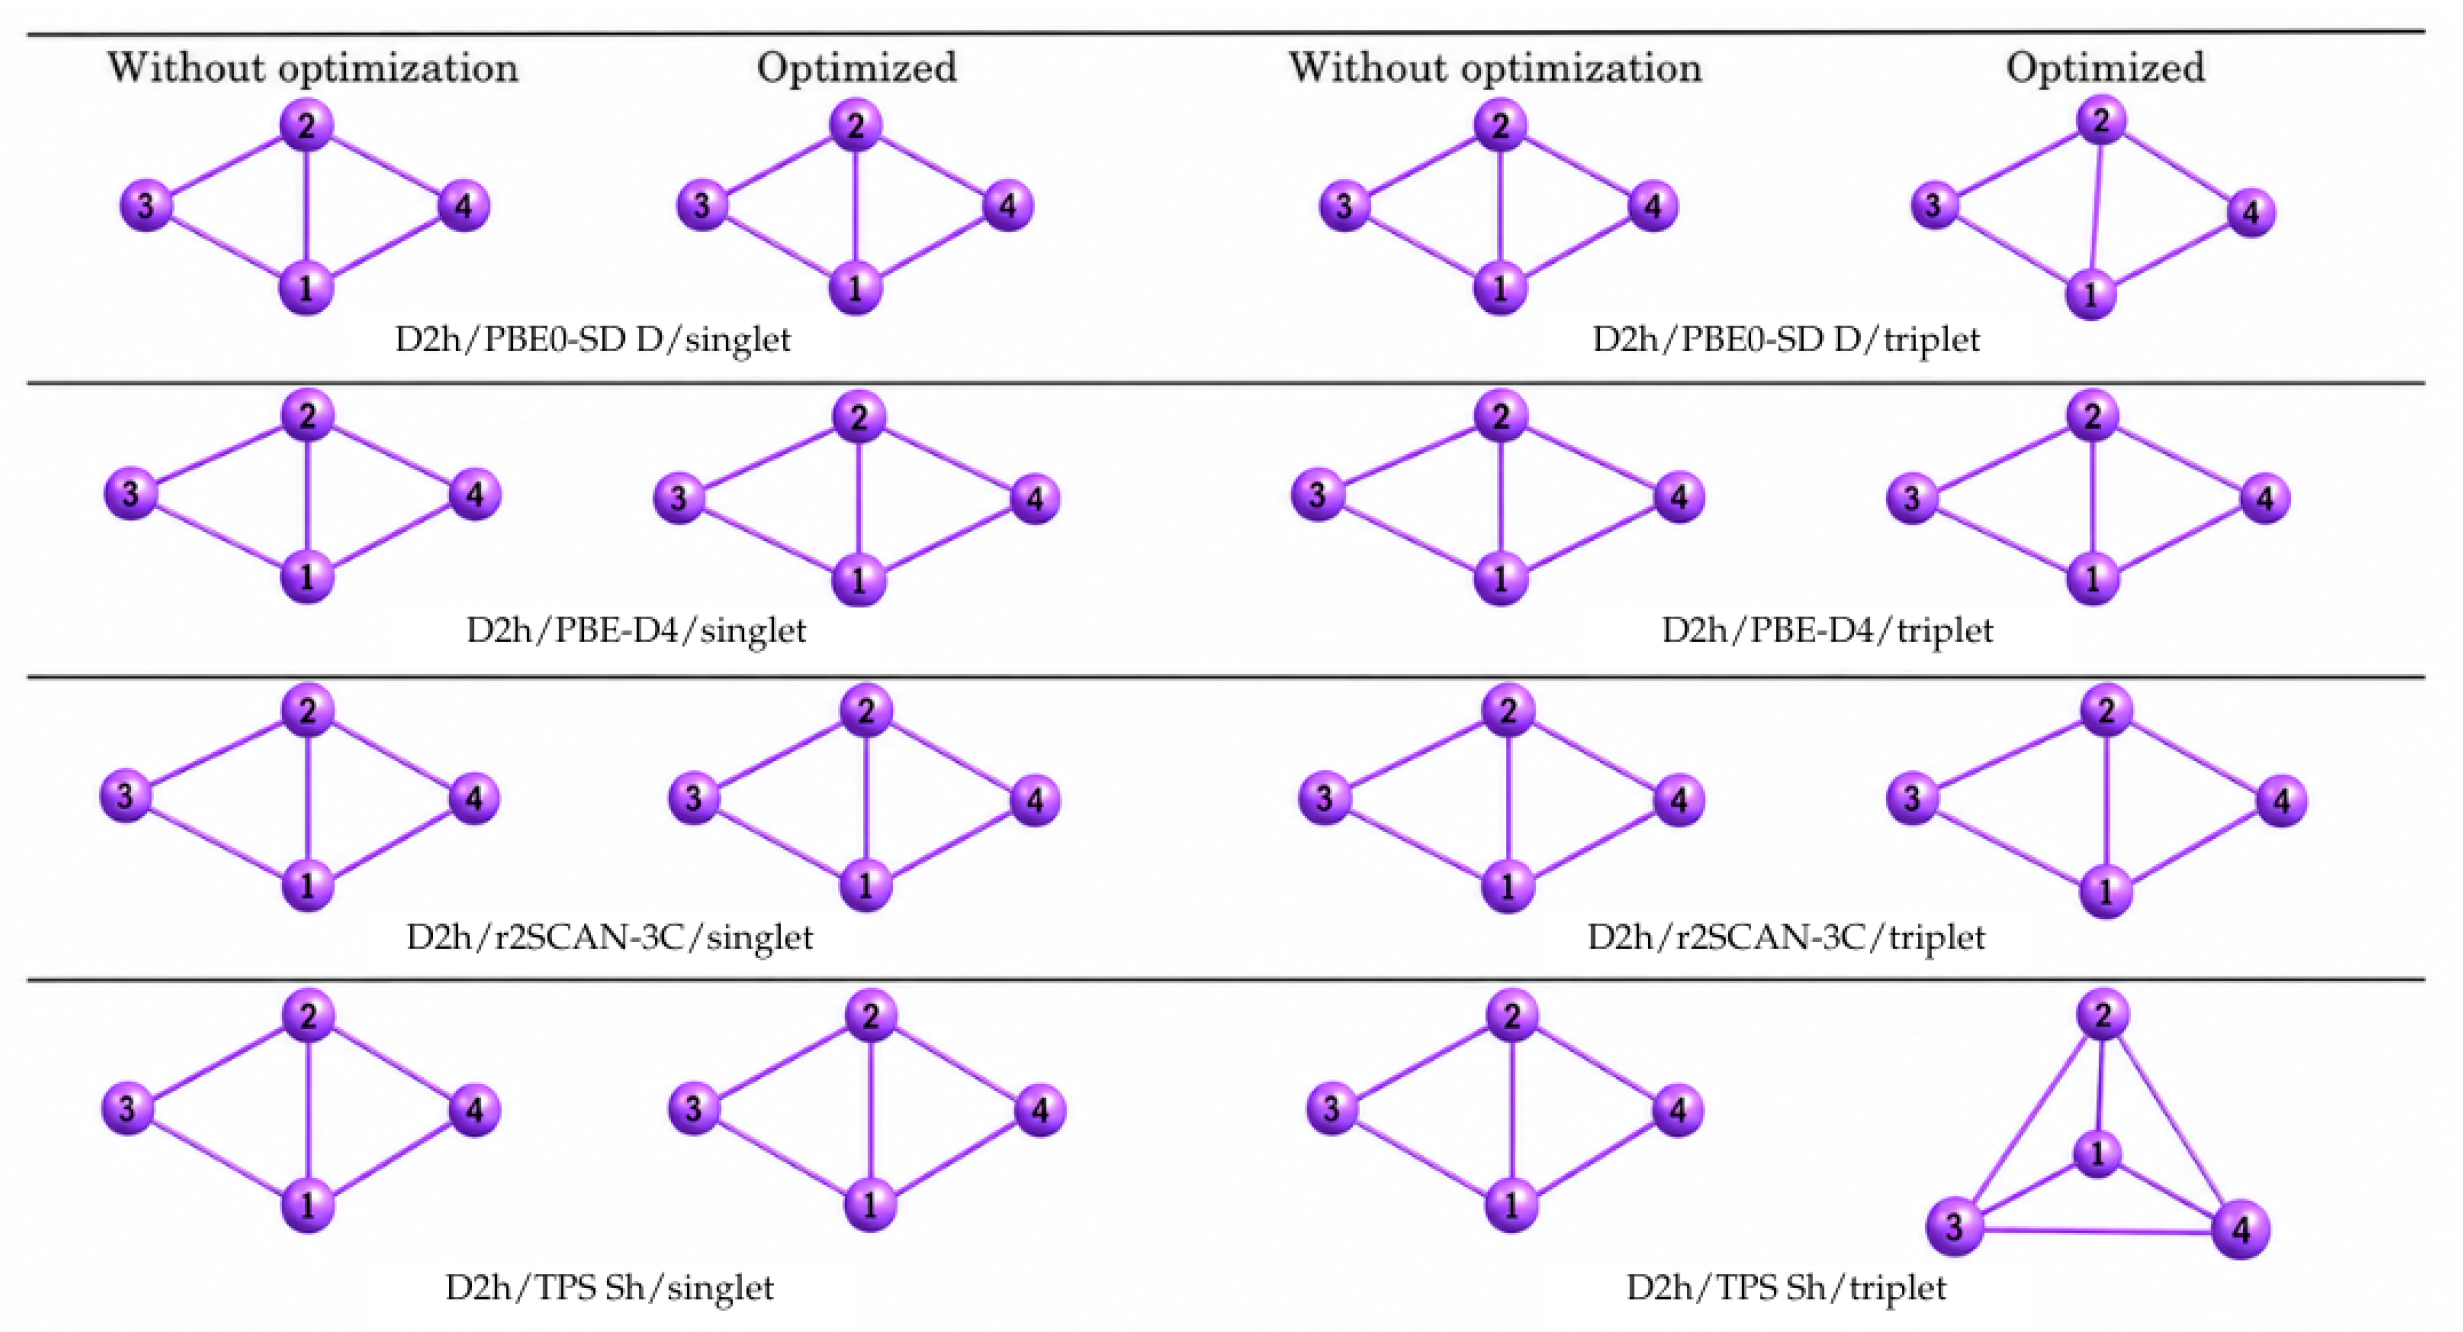
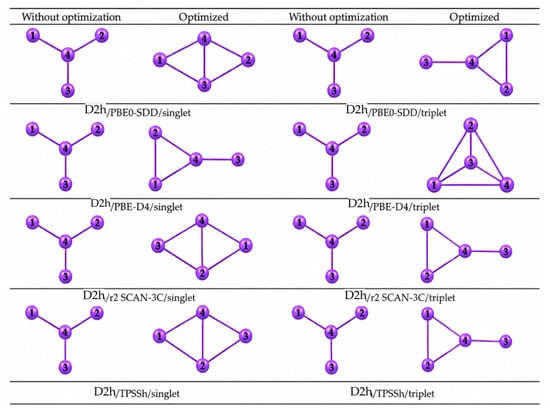
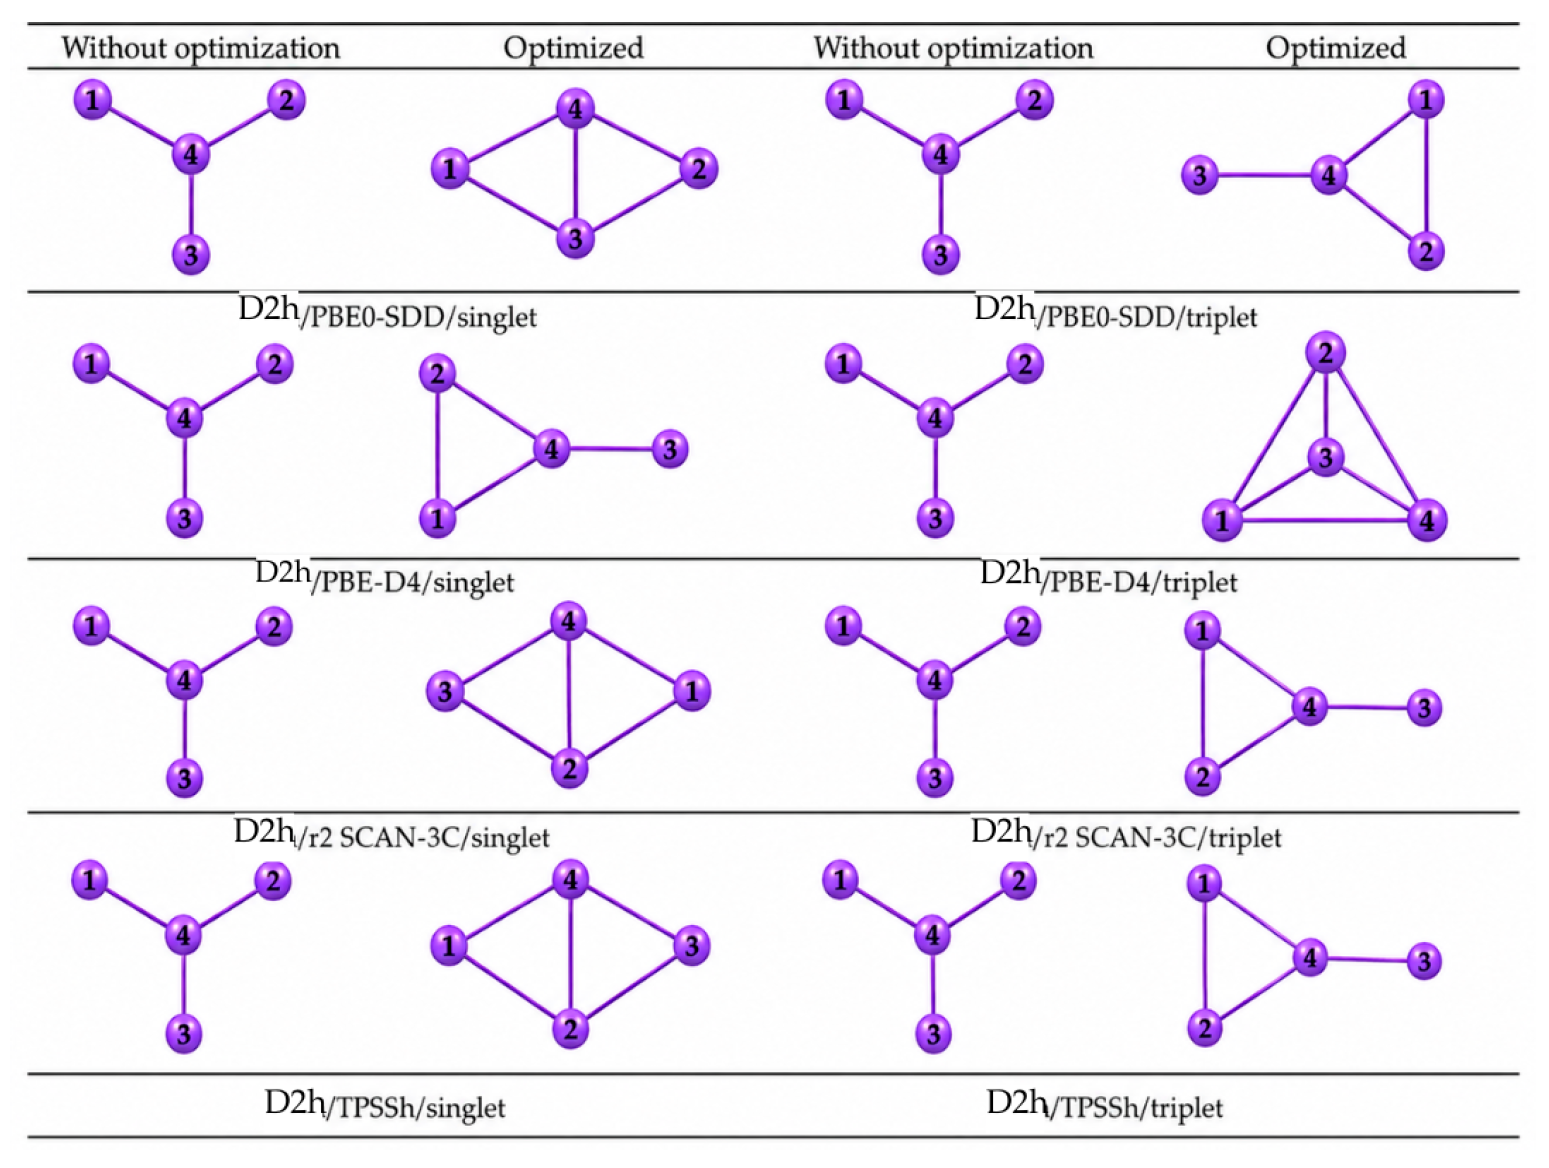
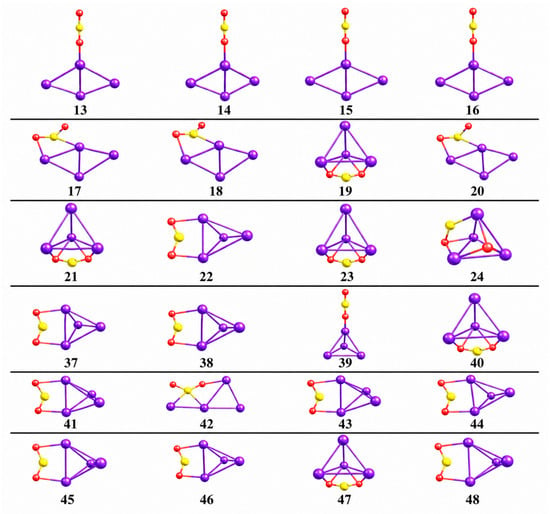
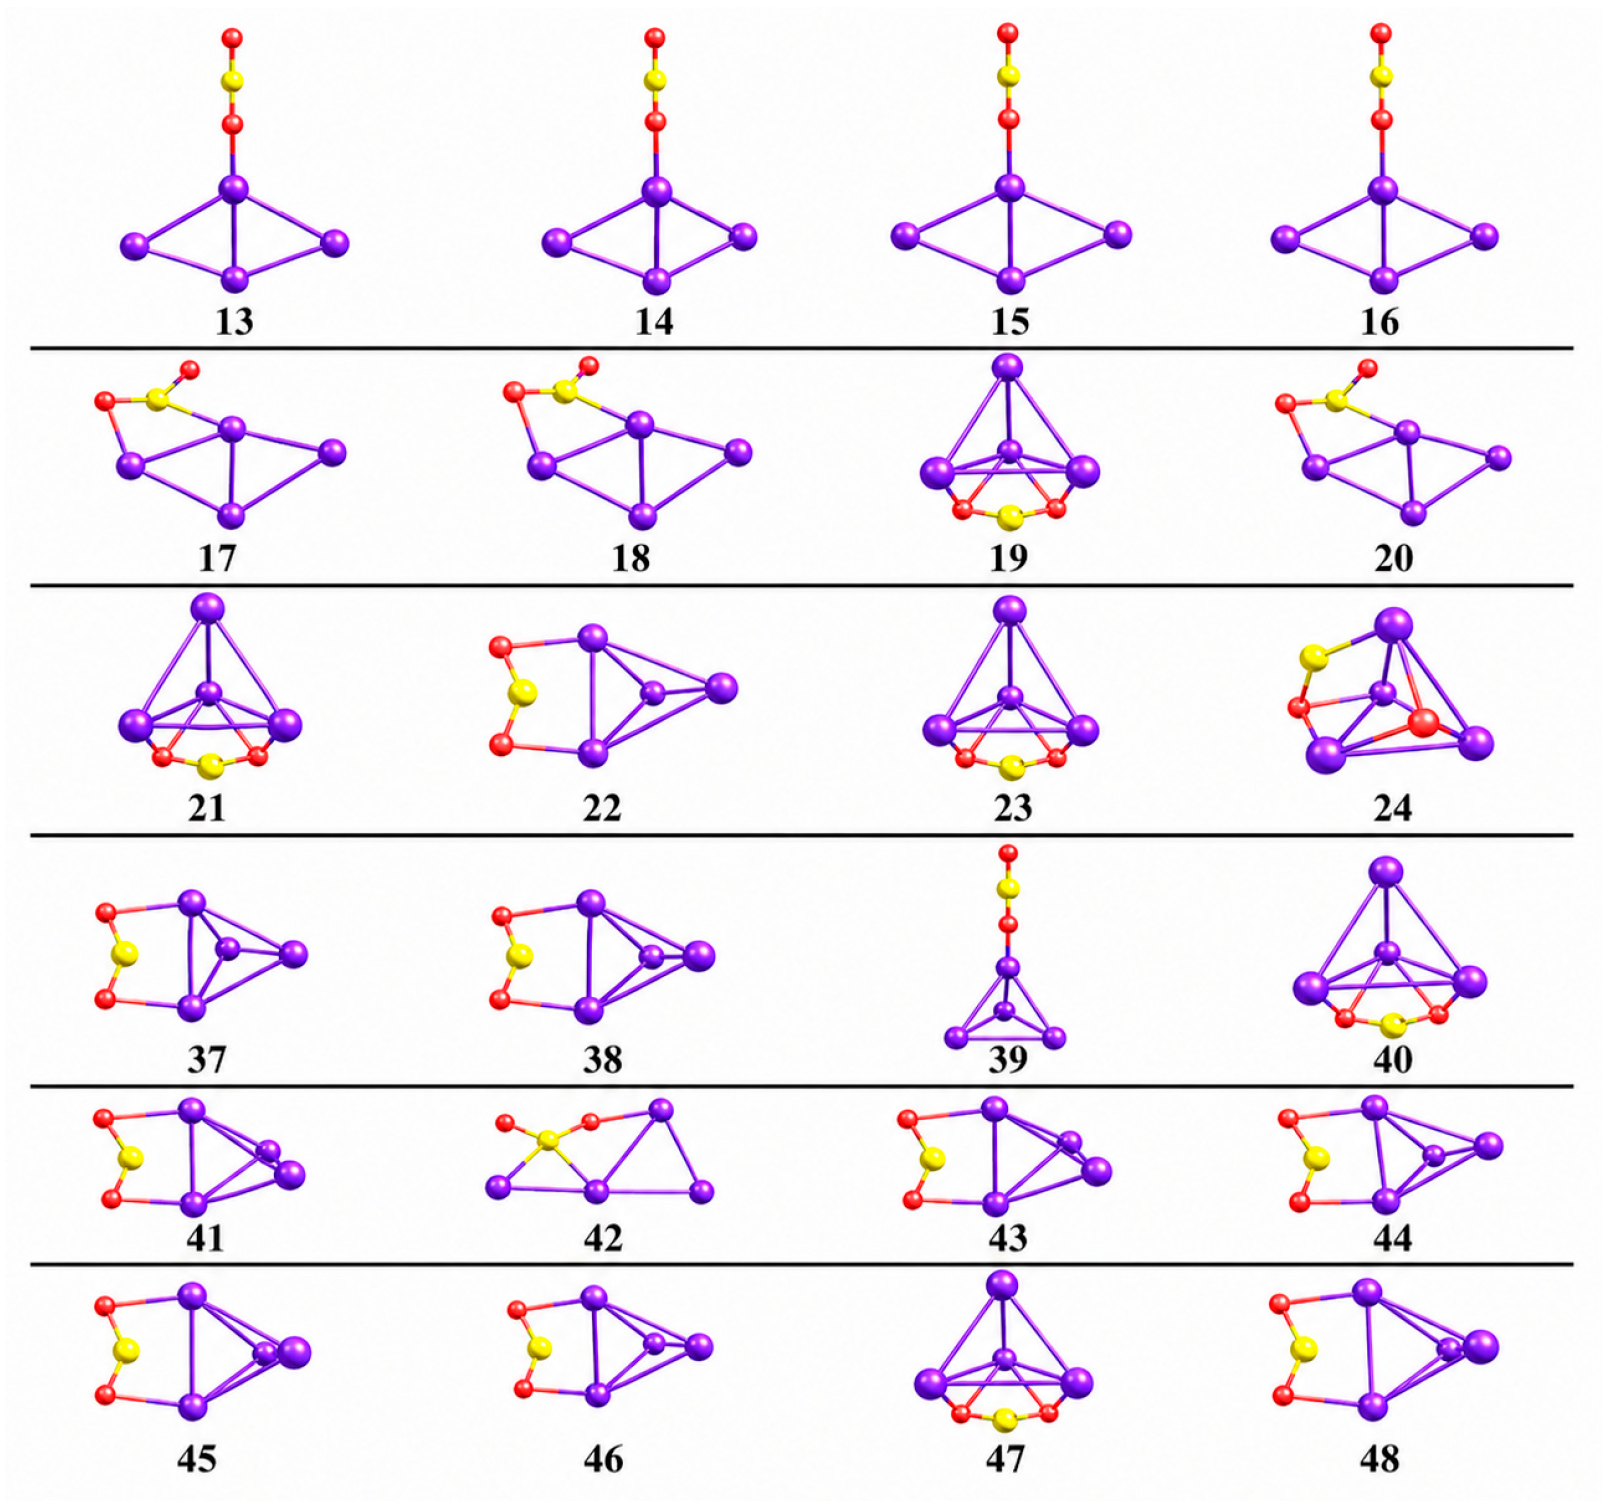
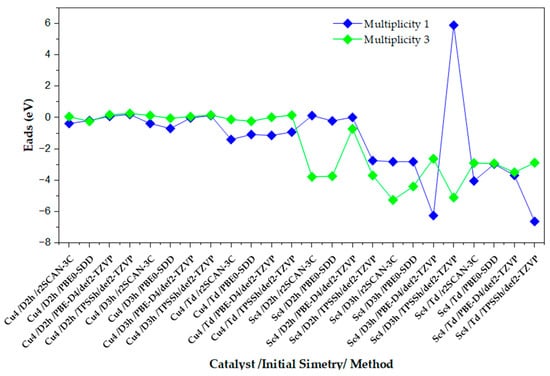
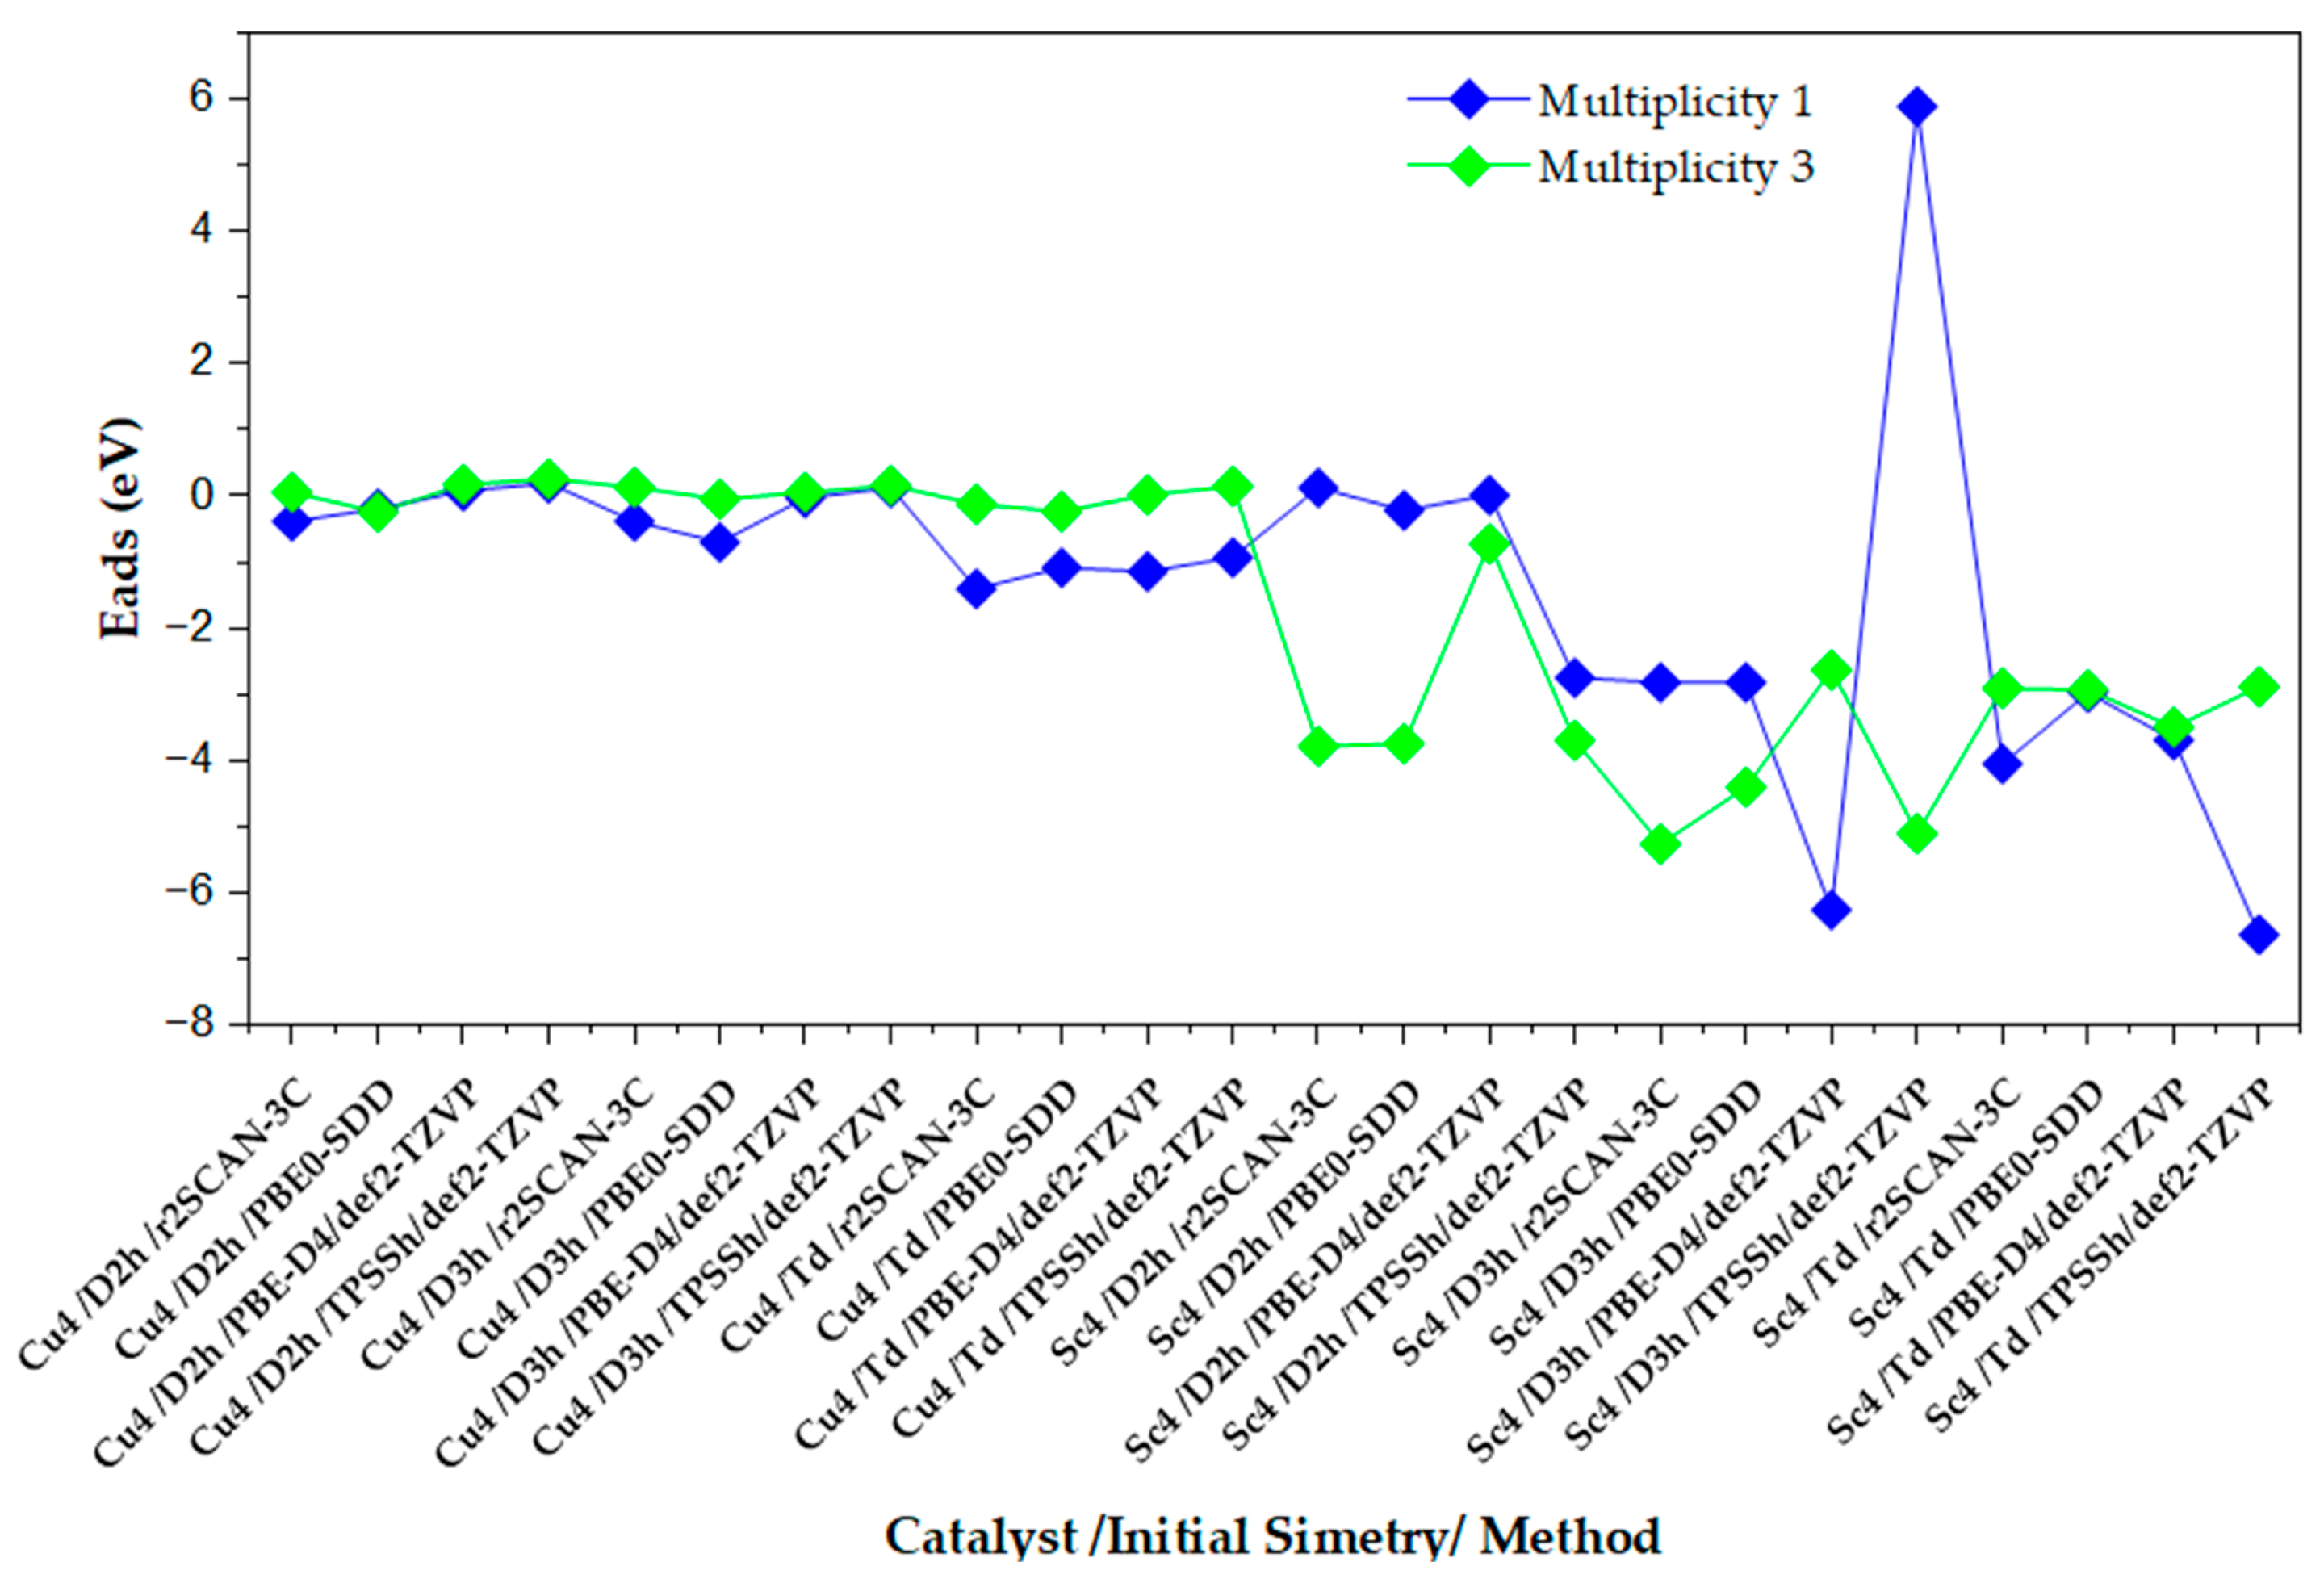
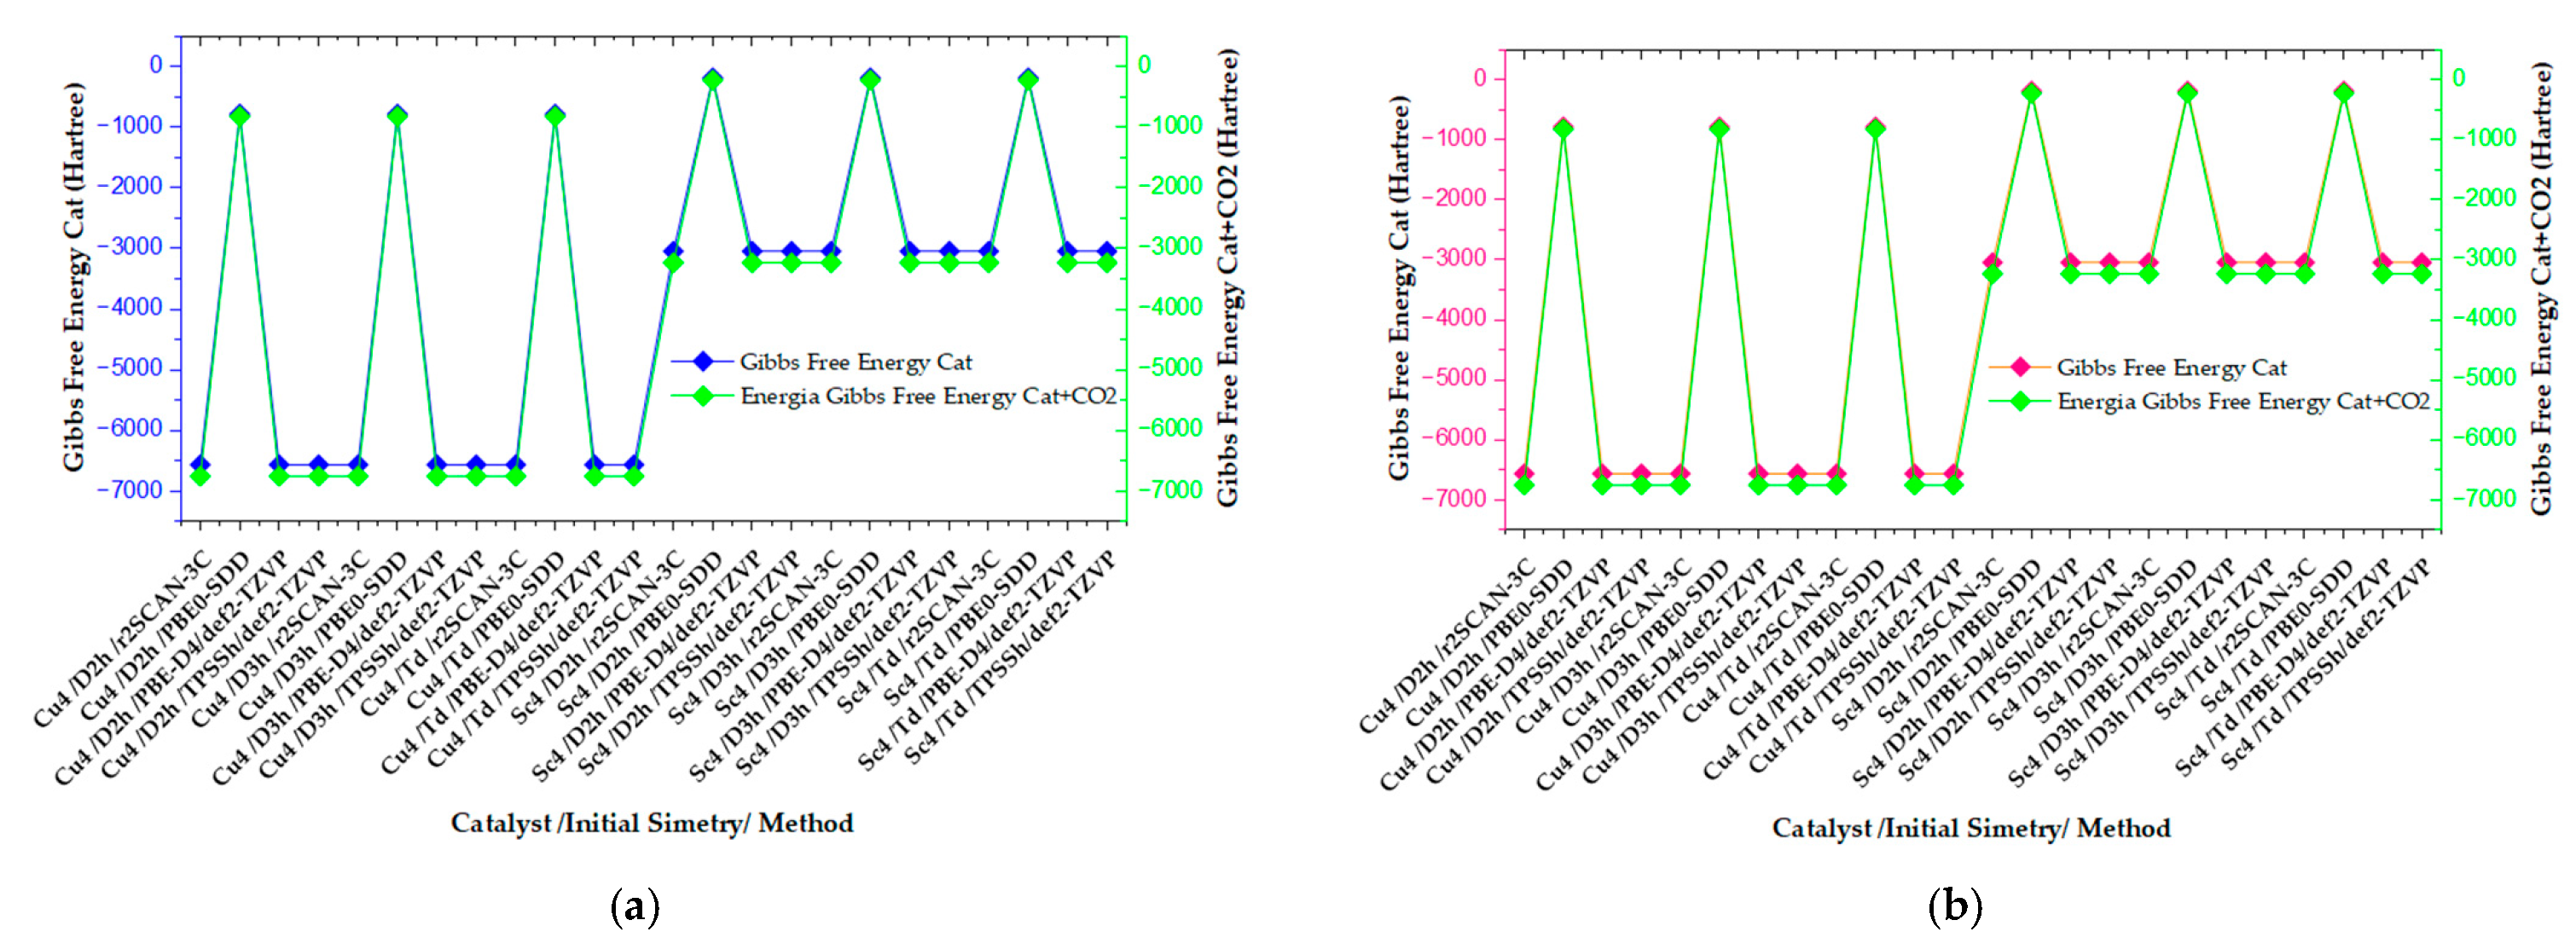

Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2026 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license.