


Abstract
The effects of NO2 and NH3 on Hg0 oxidation under fast selective catalytic reduction conditions remain unclear. To address this problem, we performed density functional theory calculations on the V2O5/TiO2(001) surface in four states: clean surface, surface covered by NH3, surface covered by NO2, and surface where species are co-adsorbed. On the clean surface, Hg0 interacts weakly with oxygen sites; the adsorption energies at terminal (Ot), bridging (Ob) and interfacial bridging (Ob′) oxygen sites are −12.31, −12.58, and −21.22 kcal mol−1, respectively. Direct oxidation by lattice oxygen encounters high activation barriers, ranging from 146.817 to 184.529 kcal mol−1. NH3 hinders the capture of Hg0. Since NH3 preferentially occupies the vanadium-related active sites, the interaction between Hg0 and the catalyst is weakened, and the adsorption energies become less negative, ranging from −11.84 to −11.04 kcal mol−1. NO2 shows the opposite effect. It strengthens Hg0 adsorption near V-centered regions and provides an alternative oxygen source for Hg0 oxidation. The NO2-assisted oxygen-transfer pathways have lower calculated barriers, ranging from 74.581 to 108.545 kcal mol−1. When NH3 and NO2 are co-adsorbed, the barrier increases to 113.326 kcal mol−1. Even so, NO2-derived oxygen provides a relatively lower-energy oxygen-transfer route than lattice oxygen among the pathways examined.
1. Introduction
NH3 selective catalytic reduction (SCR) is an efficient and mature NOx reduction technology that has been widely applied in high-temperature equipment, including coal-fired boilers, cement kilns, and municipal waste incinerators. Among the two SCR routes, the fast-SCR (FSCR) pathway is considered superior to the standard SCR (SSCR) route at lower temperatures, provided that both NO and NO2 are present in the gas mixture [1].
SSCR: 4NH3 + 4NO + O2 → 4N2 + 6H2O
FSCR: 2NH3 + NO + NO2 → 2N2 + 3H2O
In addition to NOx, flue gas from these sources contains trace mercury species [2,3]. Elemental mercury (Hg0) is particularly difficult to remove because of its high volatility, chemical inertness, and poor water solubility. As a result, it is not efficiently captured by conventional particulate-control devices or wet flue-gas desulfurization systems [4]. Catalytic oxidation of Hg0 into more soluble and easily captured oxidized mercury species is therefore regarded as a key step for deep mercury removal [5]. As SCR devices have been widely applied in combustion systems, using the same catalyst for simultaneous NOx reduction and Hg0 oxidation provides a practical solution for multi-pollutant control [6]. Therefore, vanadium-based SCR catalysts, particularly V2O5/TiO2 and V2O5–WO3/TiO2, have attracted widespread attention due to their ability to simultaneously support denitrification and mercury removal [6,7].
Experimental studies have long demonstrated that the Hg0 oxidation process over vanadium-based SCR catalysts is closely related to the surrounding SCR reaction atmosphere. Kamata et al. reported that NH3 strongly inhibits Hg0 oxidation over commercial V2O5(WO3)/TiO2 catalysts, indicating significant competition between NH3 and the surface processes involved in mercury oxidation [8]. Yang et al. subsequently discovered that the oxidation of Hg consumes surface lattice oxygen and promotes the reduction of V5+ to V4+; meanwhile, the NO conversion rate over Hg-exposed V2O5–WO3/TiO2 catalysts decreases within the 200–300 °C temperature range, further indicating a competitive relationship between Hg-related surface chemical processes and the NH3-SCR reaction [9]. In contrast, under suitable conditions, NO-derived species can promote the conversion of Hg0. Liu et al. observed that NO enhances Hg0 oxidation over commercial V2O5–WO3/TiO2 catalysts (particularly in the presence of O2), whereas excess NO inhibits Hg0 adsorption. They further proposed that NO-derived surface species, including NO2, facilitate both the adsorption and oxidation of Hg0 [10]. These early studies established that NH3 and NOx species exert markedly different effects on the chemical behavior of Hg0 over vanadium-based SCR catalysts. Recent experimental studies have further confirmed and refined these patterns. Shi et al. demonstrated that the simultaneous removal of NOx and Hg0 over V2O5/TiO2 catalysts depends heavily on the surface density and aggregation state of supported VOx species; specifically, the Hg0 oxidation rate increases as the VOx density approaches monolayer coverage, highlighting the importance of the local vanadium species environment [11]. Park et al. subsequently showed that increasing the NH3 concentration over V2O5–WO3/TiO2 catalysts enhances NO reduction but simultaneously reduces the Hg0 oxidation rate, providing direct, recent evidence of the competition between the NH3-SCR reaction and the Hg0 oxidation process [12]. Notably, Jia et al. investigated the synergistic removal of NOx and oxidation of Hg0 over V2O5–WO3/TiO2 catalysts under a simulated FSCR atmosphere, observing high Hg0 oxidation activity in the presence of NO2. Two mechanisms have been proposed: one involving surface-mediated oxidation of adsorbed Hg0, and the other a homogeneous NO2-related pathway; the latter was found to alleviate the inhibitory effect of NH3 on Hg0 oxidation [7]. Collectively, these experimental studies present a consistent qualitative picture: NH3 tends to inhibit Hg0 conversion through competitive surface interactions, whereas chemical processes derived from NO/NO2 can promote Hg0 adsorption or oxidation under suitable conditions. However, these macroscopic observations do not directly reveal how NH3 and NO2 alter the local V–O structural environment, how their pre-adsorption affects subsequent Hg0 adsorption, or how these changes influence the elementary oxygen-transfer steps involved in Hg0 oxidation. These unresolved questions regarding specific active sites represent a significant gap in our understanding of the chemical behavior of Hg0 under FSCR conditions.
Theoretical calculations have provided crucial atomic-scale insights into the adsorption and oxidation of Hg0 on supported vanadium catalysts. Suarez Negreira and Wilcox investigated Hg oxidation over vanadium–titanium (V2O5/TiO2) SCR catalysts under typical flue gas conditions, establishing an atomic-level theoretical framework to analyze the interactions between mercury species and the supported vanadium [13]. Subsequently, Gao and Li utilized a periodic V2O5/TiO2(001) model to compare Hg0 oxidation pathways, revealing the significant roles of local V–O structures, lattice oxygen, and surface redox processes in determining Hg0 reactivity [14]. Later density functional theory (DFT) studies extended this framework to modified catalyst systems. Gao et al. investigated V2O5/TiO2 catalysts modified with Cr, Fe, Co, and Nb, demonstrating that altering the local electronic and oxygen environments influences Hg0 oxidation behavior [15]. Wu et al. further examined WO3-containing V2O5/TiO2 catalysts, finding that WO3 affects surface electronic structure, mercury species adsorption, and the energetics of HCl-assisted oxidation [16]. Recent computational studies have also addressed Mo/Sb- and Ru-modified V2O5/TiO2 systems, similarly confirming that changes in local electronic structure, oxygen environment, and adsorption characteristics can significantly alter Hg0 oxidation pathways and energetics [17,18]. Despite these advances, previous theoretical research has primarily focused on the intrinsic oxidation of Hg0, HCl-assisted reaction pathways, or catalyst modification; relatively few studies have addressed the competitive effects of key FSCR components—specifically NH3 and NO2—on Hg0 adsorption and oxidation within the framework of the supported V2O5 system. This issue is particularly important because the aforementioned experimental studies indicate that NH3 inhibits while NO2 promotes Hg0 conversion, yet the atomic-scale mechanisms underlying these opposing effects remain unclear. Specifically, it remains to be elucidated how the pre-adsorption of NH3 and NO2 alters the local V–O environment, whether these species modify the surface sites associated with Hg0 adsorption or compete for them, and how their presence influences the energetics of the subsequent oxygen transfer process.
Based on a V2O5/TiO2(001) surface model, this study employs DFT calculations to investigate the atomic-scale effects of key components involved in the FSCR process on Hg0 adsorption and oxidation. Four surface environments were systematically examined: the clean surface, NH3-precovered surface, NO2-precovered surface, and NH3/NO2 co-precovered surface. By comparing optimized adsorption structures, adsorption energies, electronic properties, and characteristic oxygen-transfer pathways within a unified computational framework, we aim to elucidate how NH3 and NO2 modify the local V–O environment and the relative energetics of the Hg0 conversion process.
2. Computational Methods
2.1. Catalyst Surface Model Construction
To describe dispersed vanadia species supported on titania, a periodic V2O5/TiO2(001) surface model was constructed with reference to the benchmark studies of Gao et al. and Zhang et al. [14,19]. The model adopts the widely used anatase ad-molecule representation, which is well suited for simulating low-coverage vanadia species. In this configuration, the supported vanadia is treated as an adsorbed molecular moiety on the TiO2 substrate. Because the anatase TiO2(001) facet has been identified as one of the most catalytically active interfaces [20], it was selected as the substrate in the present work. The substrate was modeled as a two-layer anatase TiO2(001) slab with a 3 × 2 surface supercell, on which a vanadia dimer was anchored to represent the catalytically relevant V2O5 moiety. The corresponding periodic cell parameters were a = 11.328 Å, b = 7.552 Å, and c = 20.529 Å, with α = β = γ = 90°. The periodic cell contains 12 Ti, 29 O, and 2 V atoms (Ti12V2O29), corresponding stoichiometrically to a Ti12O24 slab with a supported V2O5 species. This simplified V2O5/TiO2(001) surface was adopted as a controlled model to isolate the effects of the selected FSCR components on the local V–O environment and Hg0 conversion.
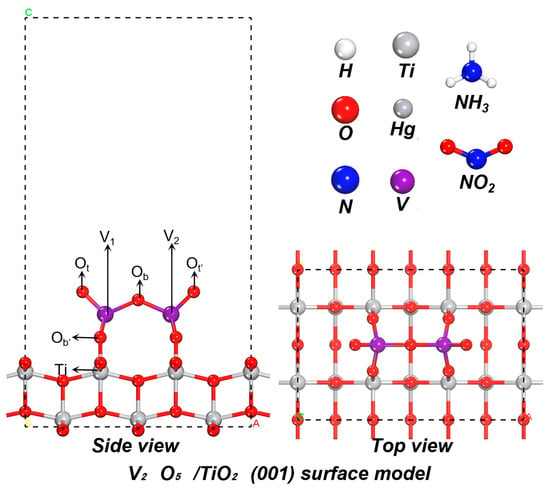
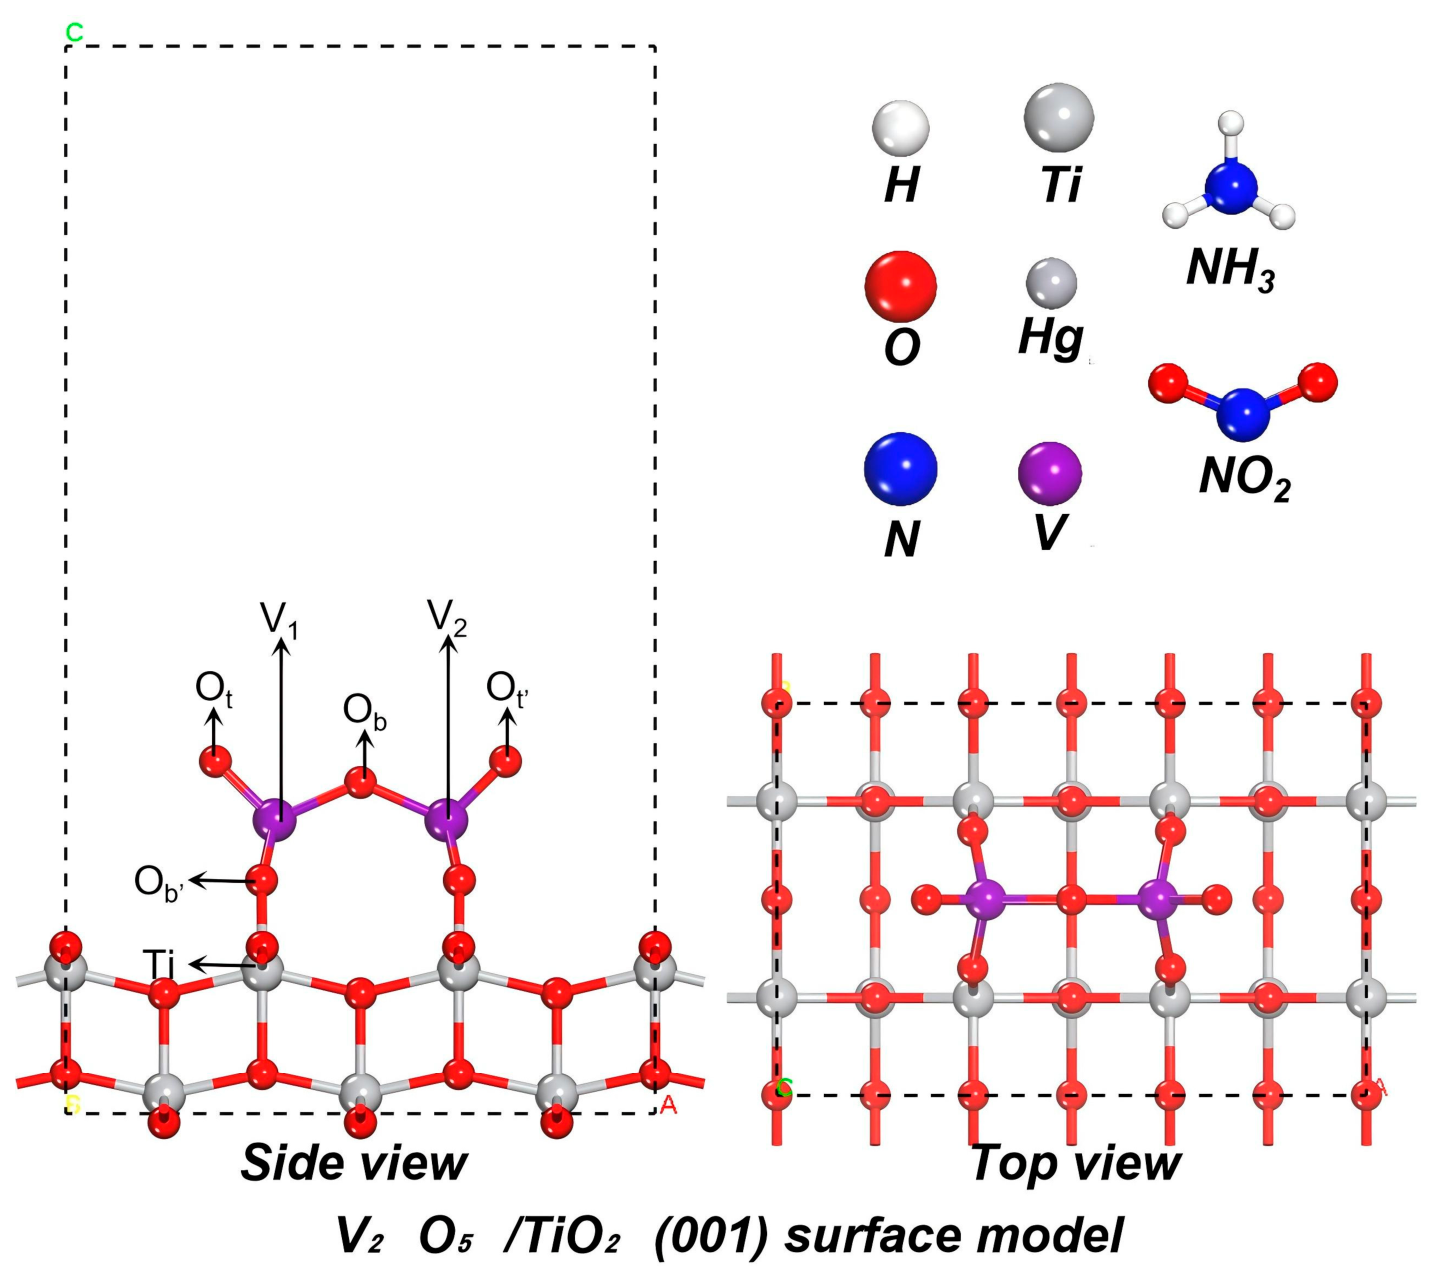
Motivated by experimental studies showing that NH3 and NO2 in FSCR atmospheres can significantly influence Hg0 oxidation over vanadia-based SCR catalysts [7], the calculations were organized in a stepwise manner to clarify the individual and combined effects of these species on Hg0 conversion over V2O5/TiO2(001). First, Hg0 adsorption and oxidation on the clean surface were investigated to establish a reference. Next, the adsorption behaviors of NH3 and NO2 were examined separately, followed by Hg0 adsorption and oxidation on the corresponding precovered surfaces. Finally, NH3/NO2 co-precovered surfaces were considered to evaluate their combined influence on Hg0 adsorption and oxidation. For each stage, multiple initial configurations were examined to identify representative optimized structures and possible reaction precursors. The resulting adsorption structures were subsequently used for transition-state calculations and comparison of the corresponding oxidation pathways. The computational surface model used throughout this workflow is shown in Figure 1.
Figure 1.
Computational model of the V2O5/TiO2(001) surface used for DFT simulations.
The oxygen atoms in the supported vanadia species can be divided into three types according to their local coordination environments, i.e., terminal oxygen (Ot), bridging oxygen in the V–O–V linkage (Ob), and interfacial bridging oxygen in the V–O–Ti linkage (Ob′). This classification serves as a useful structural framework for identifying potential adsorption sites involved in Hg0 oxidation under FSCR conditions.
To assess the reliability of the adopted surface model, the optimized bond lengths of the V2O5/TiO2(001) structure were compared with previously reported theoretical and experimental values, as listed in Table 1. The calculated bond lengths are in good agreement with the literature data, with only small deviations, indicating that the present model is suitable for the subsequent calculations.
Table 1.
Comparison of characteristic bond lengths in the optimized V2O5/TiO2(001) model.
2.2. DFT Calculation Details
First-principles calculations were performed using the DMol3 module implemented in BIOVIA Materials Studio 2017 (Dassault Systèmes, San Diego, CA, USA) [23]. To account for the open-shell nature of the gas-phase species involved in the FSCR system, spin-unrestricted calculations were employed throughout, with formal spin used as the initial state to facilitate self-consistent field (SCF) convergence. Following previous studies, exchange–correlation interactions were described using the generalized gradient approximation (GGA) with the Perdew–Burke–Ernzerhof (PBE) functional, which has been widely used in previous DMol3 studies of V2O5/TiO2-based SCR catalysts and Hg0 oxidation pathways [16,24]. A double-numerical basis set with polarization functions (DNP) was used to expand the valence electron wave functions. To account for relativistic effects, the core electrons of vanadium were treated using density functional semi-core pseudopotentials (DSPP). In addition, the Tkatchenko–Scheffler (TS) dispersion correction was included to describe non-covalent van der Waals (vdW) interactions between adsorbates and the catalyst surface. To improve the description of localized V 3d states involved in the redox chemistry of the supported vanadia species, a Hubbard U correction of 4.0 eV was applied to the V 3d orbitals throughout the calculations [25]. The real-space global orbital cutoff radius was set to 5.2 Å, and a thermal smearing of 0.01 Ha was applied to the orbital occupations.
Geometry optimization was considered converged when the changes in total energy, maximum force, and maximum displacement were below 5.0 × 10⁻⁵ Ha, 2.0 × 10⁻² Ha Å−1, and 5.0 × 10⁻² Å, respectively. Brillouin-zone sampling was performed using Monkhorst–Pack k-point meshes of 3 × 3 × 3 for bulk anatase TiO2 and 3 × 3 × 1 for the surface slab models. To avoid spurious interactions between periodic images, a vacuum layer of 15 Å was introduced along the z direction.
2.3. Adsorption and Transition-State Calculations
For adsorption calculations, the atoms in the bottom layer of the TiO2(001) slab were fixed, whereas the upper-layer atoms, the supported V2O5 species, and the adsorbed molecules were fully relaxed. The adsorption strength of each gaseous species on the catalyst surface was evaluated using the adsorption energy, Eads, defined as
where Eadsorbate/substrate is the total energy of the adsorbate–surface complex, Eadsorbate is the energy of the isolated adsorbate, and Esubstrate is the energy of the clean surface. A more negative Eads value indicates stronger adsorption.
Eads = Eadsorbate/substrate − Eadsorbate − Esubstrate
To map the elementary reaction pathways for Hg0 oxidation under FSCR conditions, transition-state (TS) searches were performed using the complete linear synchronous transit/quadratic synchronous transit (LST/QST) method. All optimized stationary points were further verified by vibrational frequency analysis. A stable intermediate was confirmed by the absence of imaginary frequencies, whereas a valid transition state was identified by the presence of a single imaginary frequency along the reaction coordinate. The activation barrier, Ea, for each elementary step was calculated as
where ETS and EIM denote the energies of the transition state and the corresponding adsorbed intermediate, respectively.
Ea = ETS − EIM
Since thermal and entropic corrections were not included, the calculated adsorption energies and activation barriers are interpreted primarily in terms of qualitative energetic trends among the different pathways within the present computational framework.
3. Results and Discussion
3.1. Hg0 Adsorption and Oxidation on V2O5/TiO2(001)
3.1.1. Hg0 Adsorption Behavior
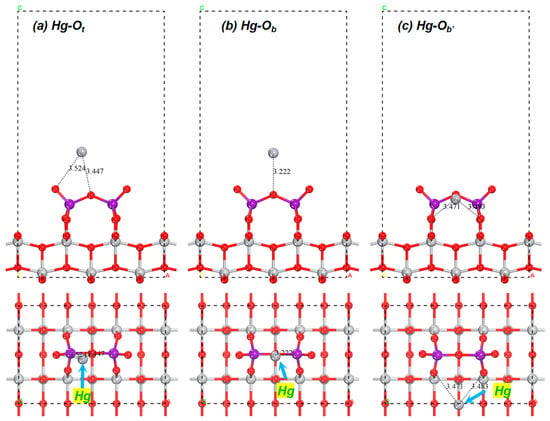
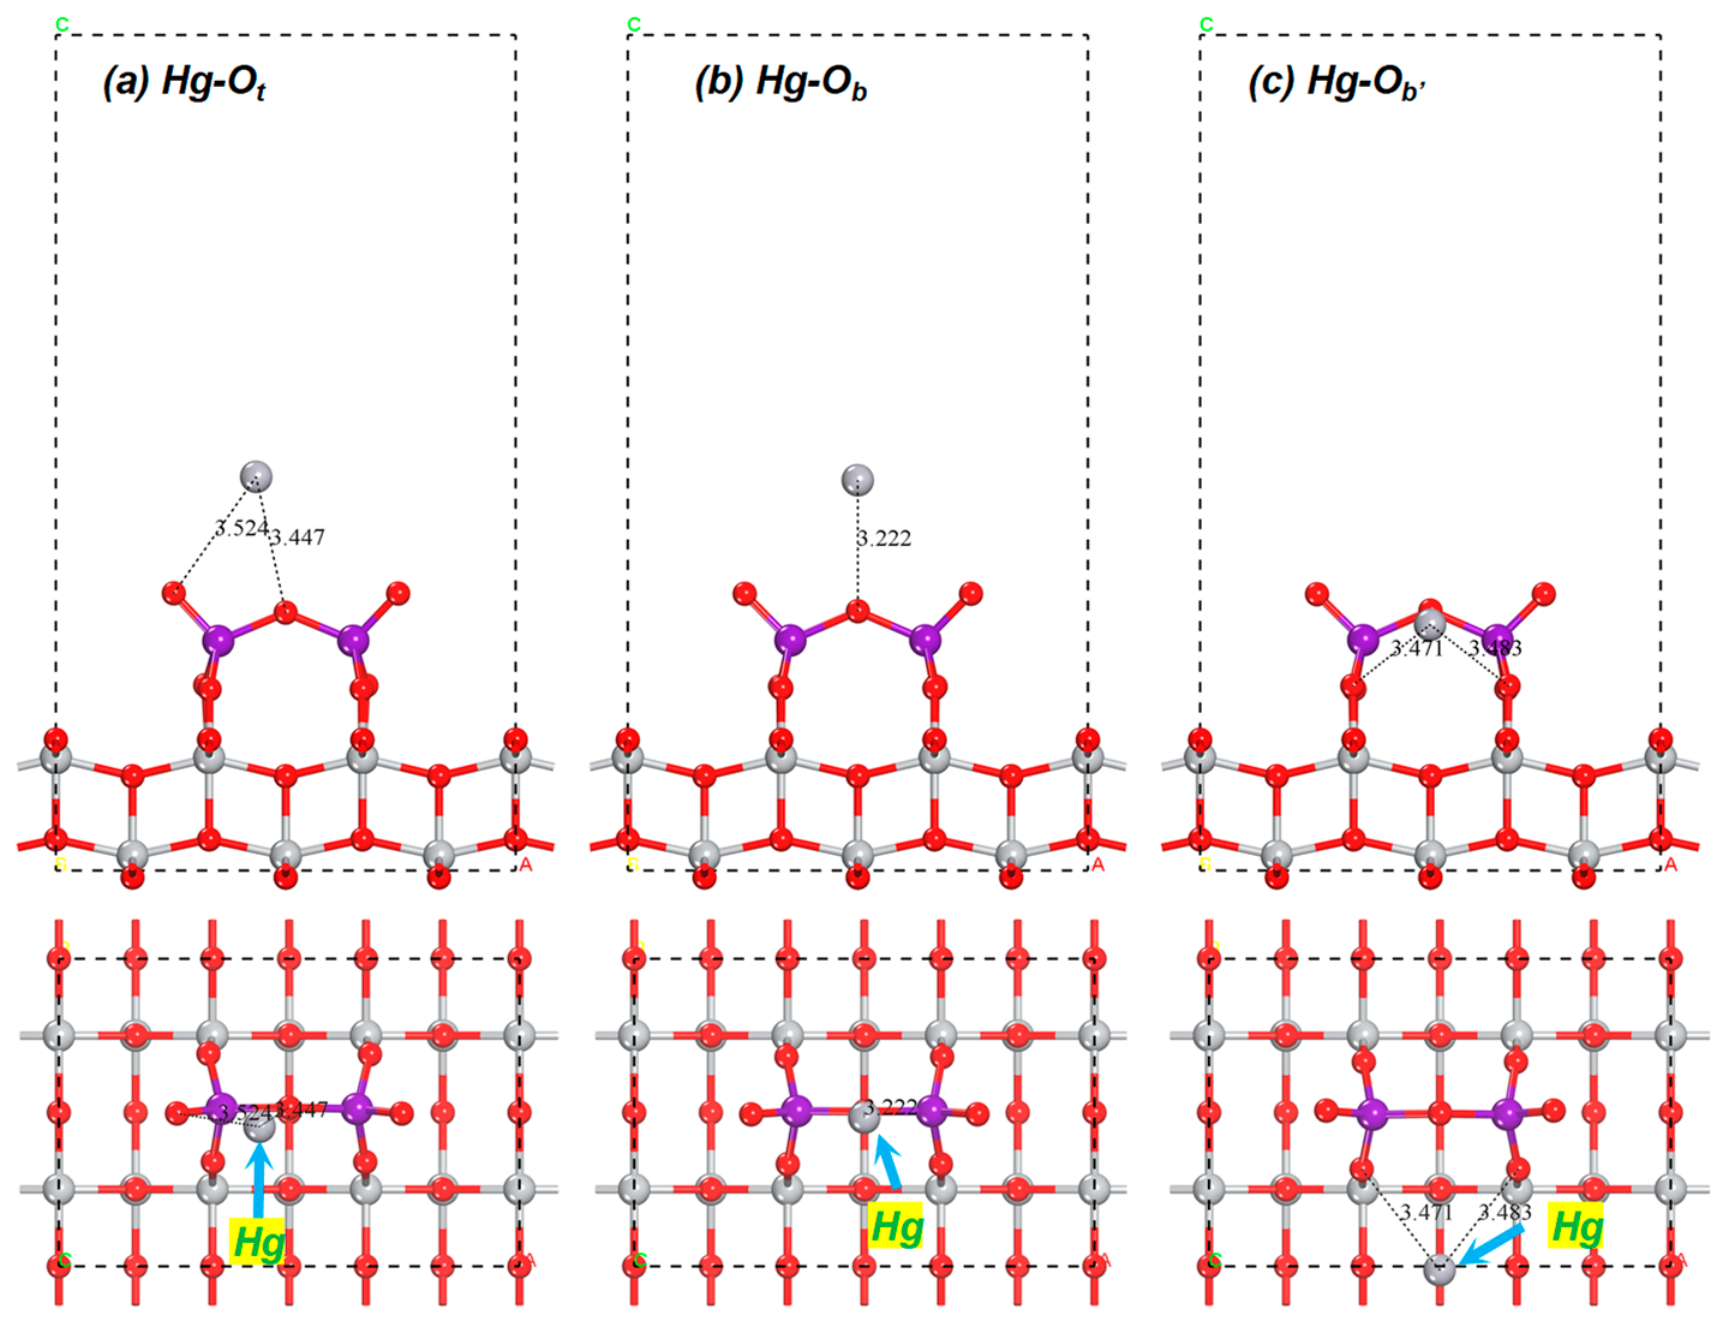
We first explored the adsorption and oxidation behavior of Hg on the clean V2O5/TiO2 (001) surface. In this section, the configurations are named based on the following principles: for example, Hg-Ot means that Hg is initially placed above the Ot site and then geometrically optimized. The optimized structures are shown in Figure 2, and the corresponding adsorption energies, structural parameters and Mulliken charges are shown in Table 2.
Figure 2.
Optimized configurations of Hg0 adsorbed at different oxygen sites on V2O5/TiO2(001).
Table 2.
Adsorption energies, Hg–O distances, and Mulliken charges for Hg0 adsorption on V2O5/TiO2(001).
During the structural relaxation, the Hg-Ob′ configuration exhibits the most negative adsorption energy (−21.22 kcal mol−1), which is significantly lower than those of Hg-Ot (−12.31 kcal mol−1) and Hg-Ob (−12.58 kcal mol−1). This indicates that the interface Ob′ site associated with the V-O-Ti bond is the most favorable region for Hg adsorption. In contrast, the similar adsorption energies of Hg-Ot and Hg-Ob indicate that the adsorption stability of Hg0 at the terminal oxygen and V-O-V bridge oxygen sites is relatively weak. The optimized geometric structure reveals further insights. In the Hg-Ot model, Hg moves above the vanadium dimer and stabilizes between two adjacent oxygen atoms, with Hg-O distances of 3.524 and 3.447 Å, respectively. In the Hg-Ob configuration, Hg0 is located above the central bridge oxygen of the V-O-V unit, showing the shortest Hg-O distance (3.222 Å). For the Hg-Ob′ configuration, Hg migrates to the oxide-support interface and is stabilized between two adjacent interface oxygen atoms, with Hg-O distances of 3.471 and 3.483 Å, respectively. Despite these structural differences, all the Hg-O distances are still relatively long, indicating that no strong Hg–O bond is formed. Interestingly, although Hg-Ob′ does not show the shortest bond length, it shows the strongest adsorption. This indicates that the adsorption stability is affected not only by geometric factors but also by the electronic properties of the adsorption sites, especially at the oxide-support interface.
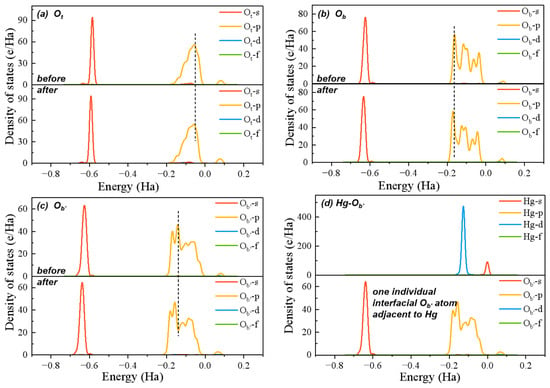
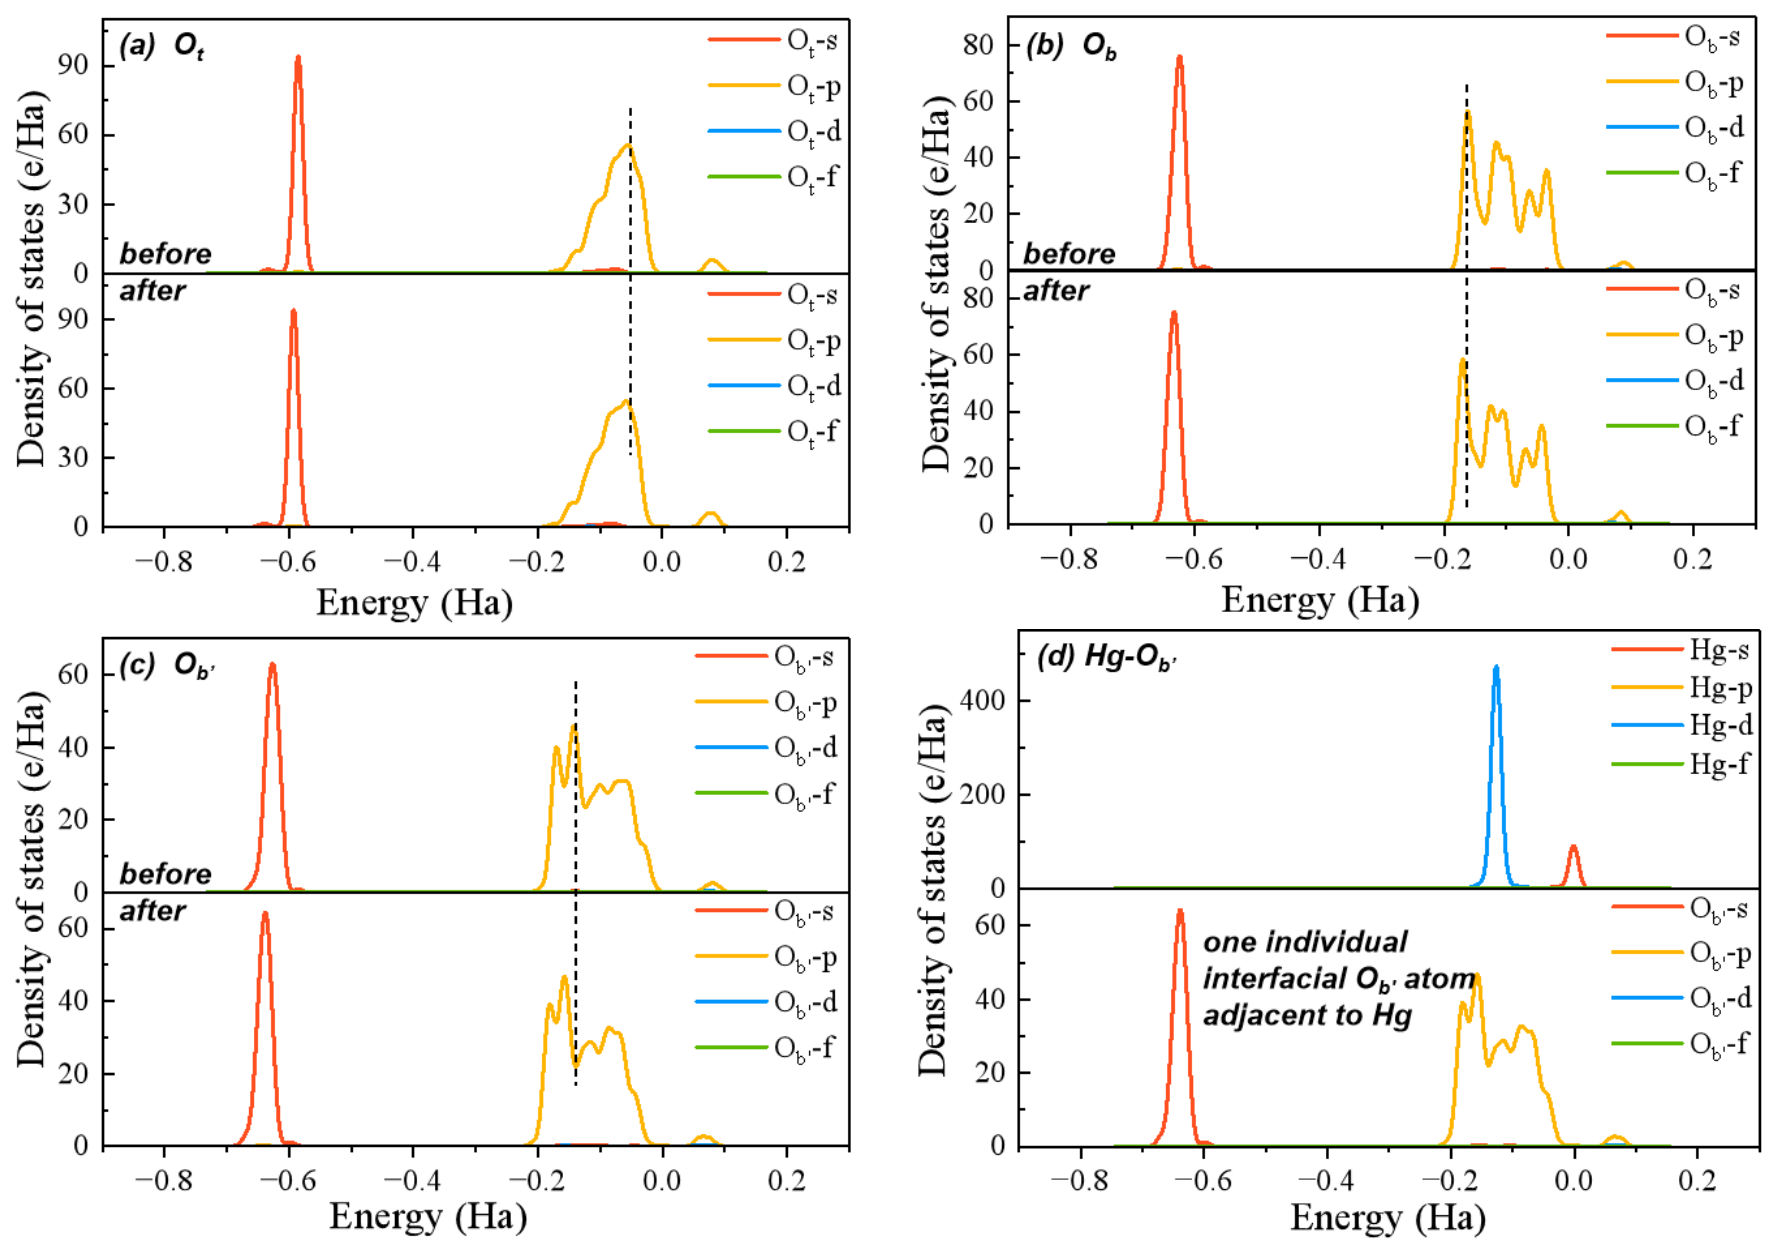
Mulliken charge analysis further supports this explanation. The charge of Hg in Hg-Ot, Hg-Ob and Hg-Ob′ configurations is 0.113 e, 0.115 e and 0.167 e, respectively. The small charge transfer in all configurations indicates that Hg basically maintains neutral characteristics after adsorption, which is consistent with the weak interaction inferred from the long Hg-O distance. However, the relatively high positive charge in the Hg-Ob′ configuration indicates a stronger electron perturbation at the interface V-O-Ti site. Additional insights come from the partial density of states (PDOS) diagram shown in Figure 3. For Hg-Ot and Hg-Ob, the PDOS of the corresponding oxygen atoms shows little change, and the peak shape, intensity and energy distribution exhibit almost no change. This weak electronic response is consistent with the small charge transfer and similar adsorption energy observed in these configurations. In contrast, the Hg-Ob′ system undergoes more significant electronic changes. After Hg adsorption, the p state of the interface Ob′ atom moves towards lower energy, reflecting a stronger local electronic structure perturbation. There is obvious hybridization between the d state of Hg and the p state of Ob′, whereas this orbital coupling is much weaker in Hg-Ot and Hg0-Ob. These electronic structure features explain the stronger adsorption of Hg at the Ob′ site. In other words, the higher stability is mainly due to the interfacial electron interaction, especially the Hg(d)-Ob′(p) coupling. Combined with the larger Mulliken charge on the Hg0 atom in the Hg-Ob′ configuration, the PDOS results show that the V-O-Ti interface region provides the most favorable local environment for the electron perturbation and adsorption stability of Hg on the V2O5/TiO2(001) surface.
Figure 3.
Local atom- and orbital-resolved PDOS for Hg0 adsorption on V2O5/TiO2(001), focusing on Hg and the surface O atom directly involved in each adsorption environment.
3.1.2. Hg0 Oxidative Desorption Behavior
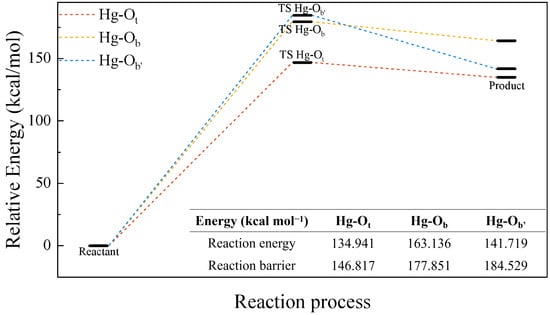
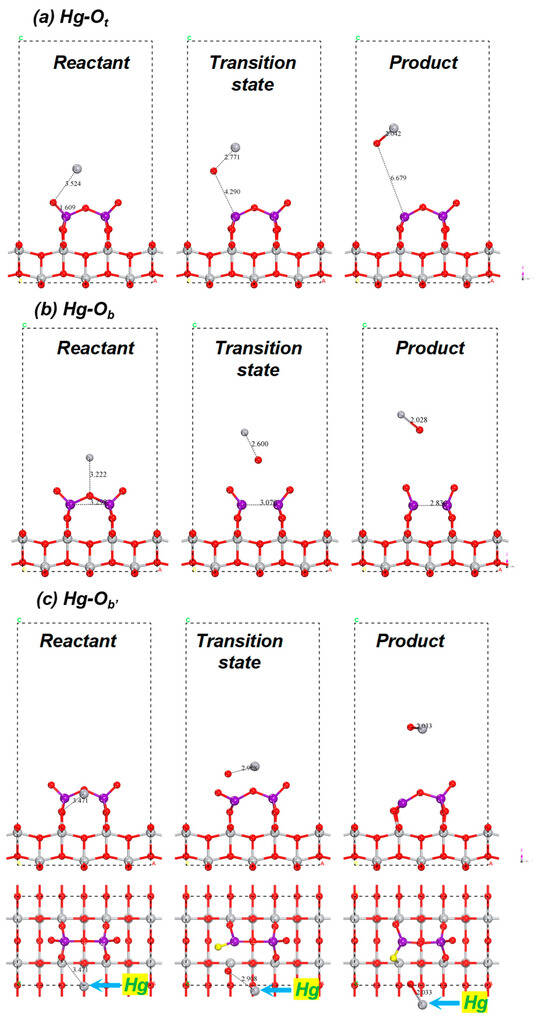
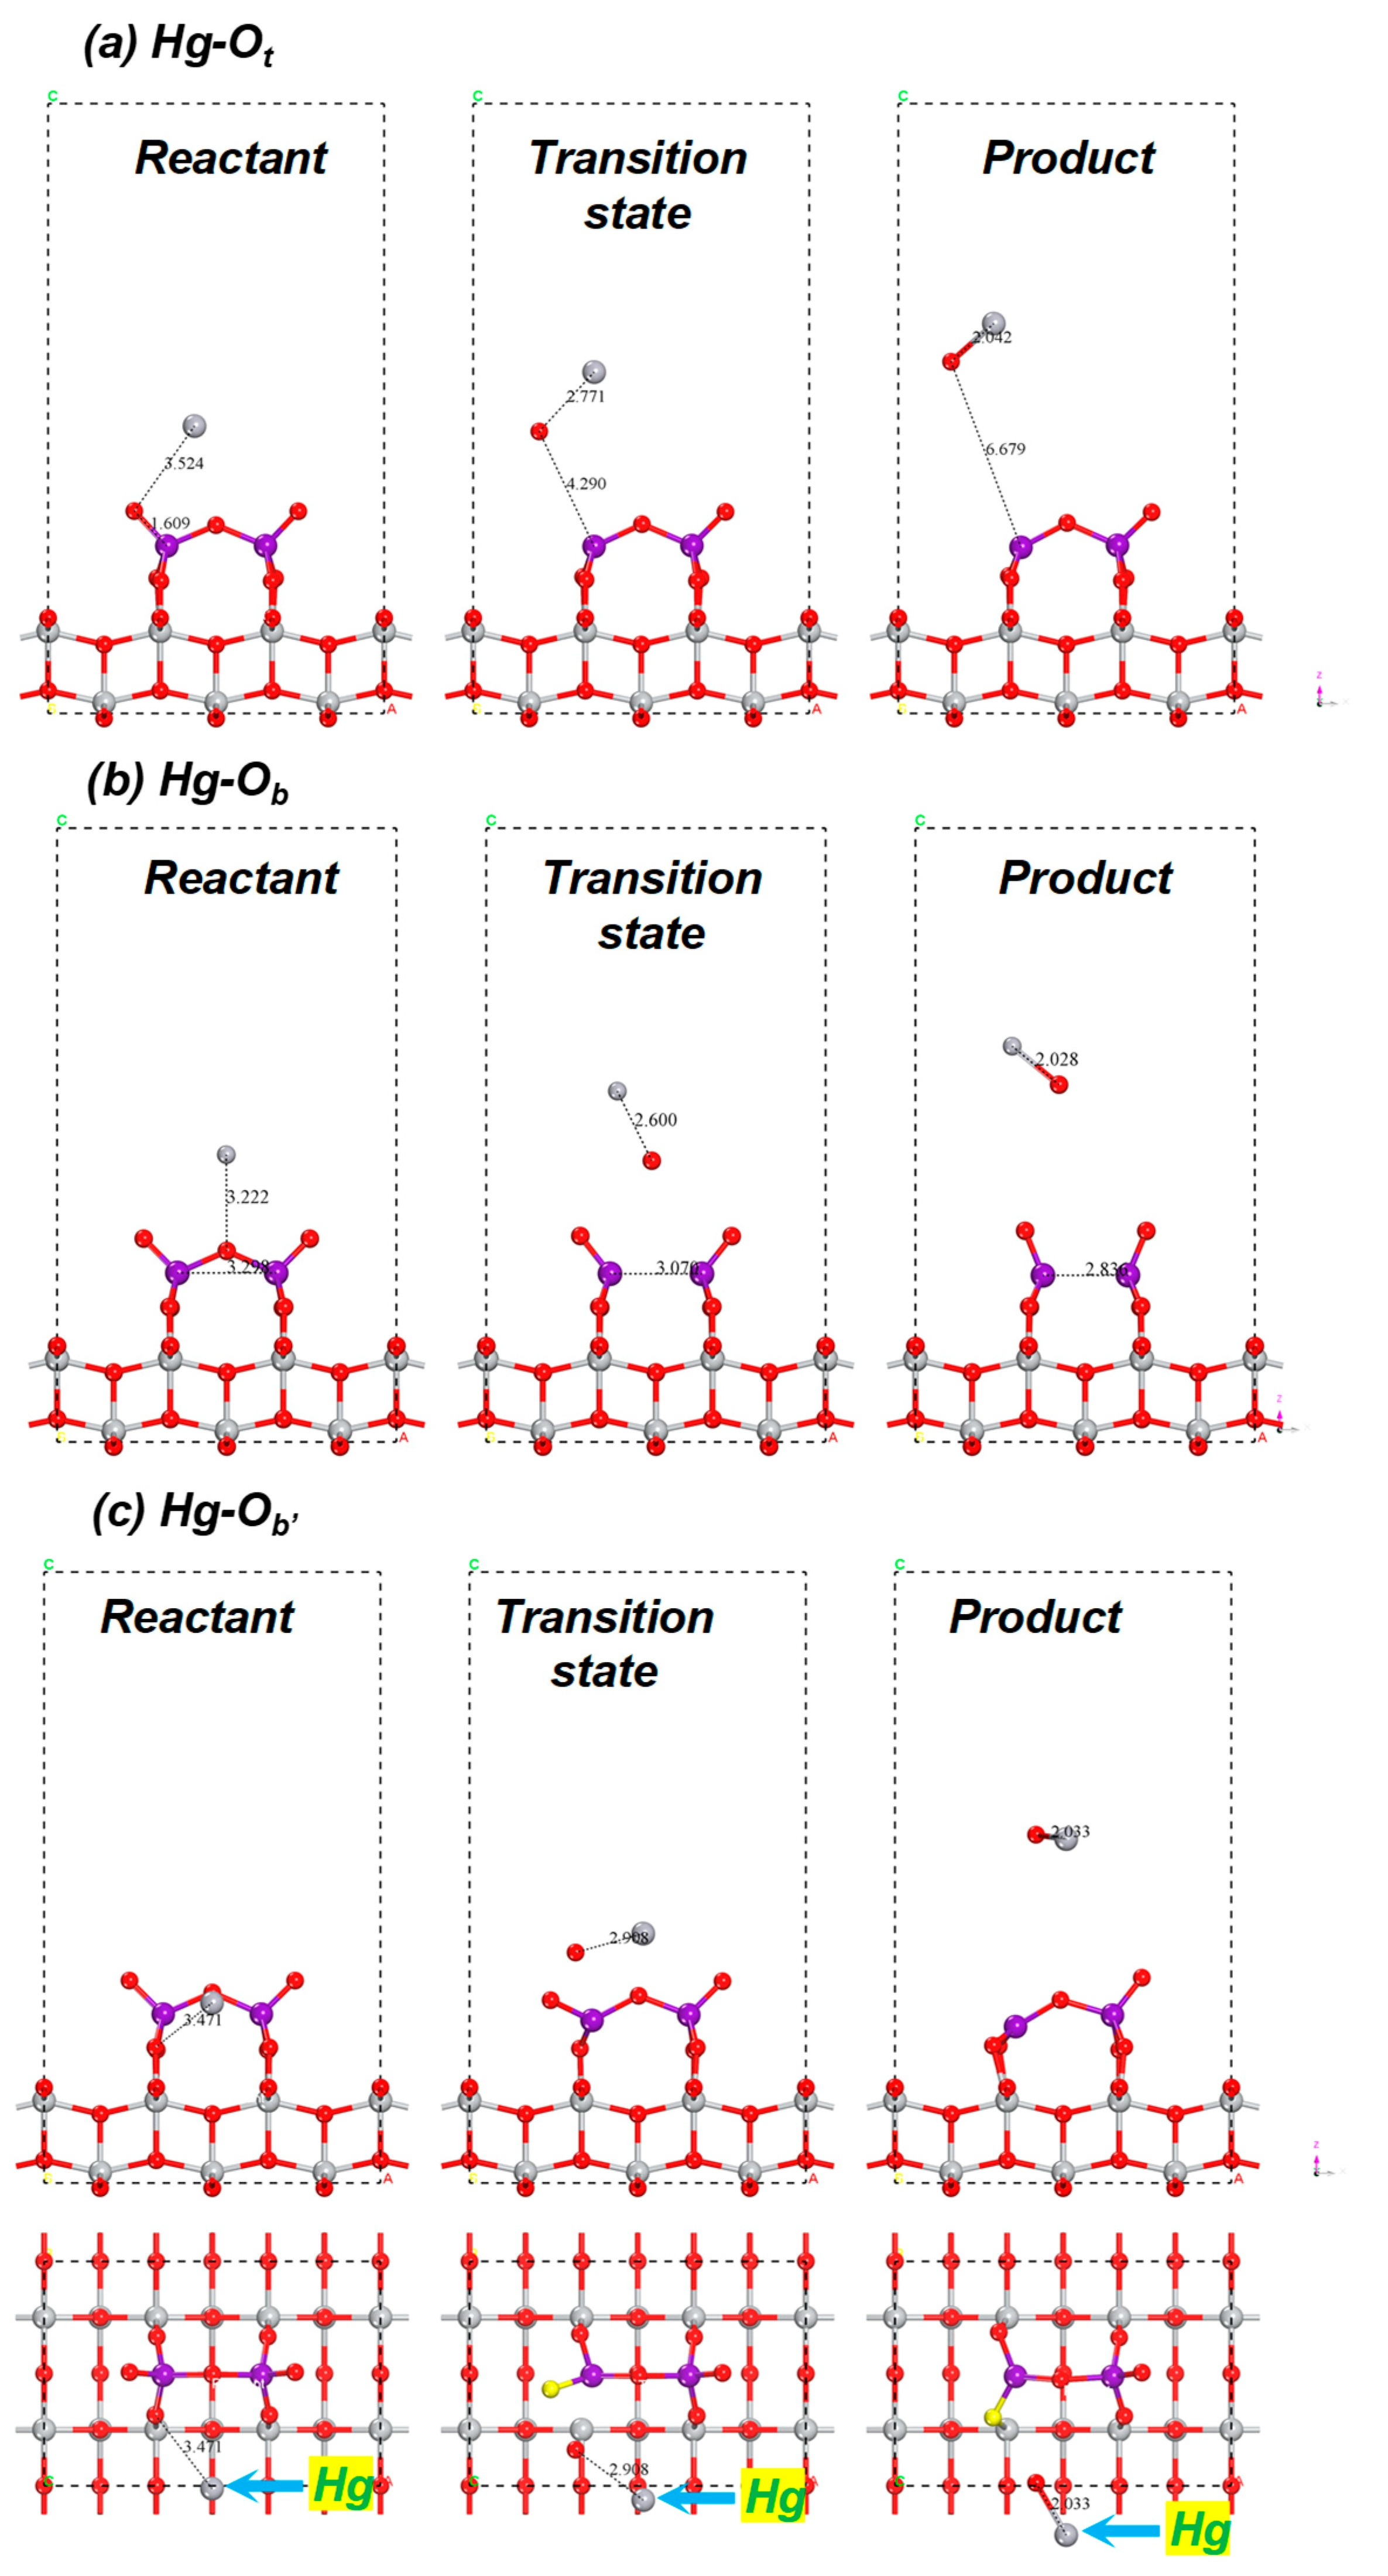
Transition-state calculations were carried out to compare Hg0 oxidation and oxidative desorption at the Ot, Ob, and Ob′ sites on V2O5/TiO2(001). The energy profiles are shown in Figure 4, and the optimized structures along the reaction pathways are presented in Figure 5. All three pathways are strongly endothermic and exhibit high calculated activation barriers, indicating that direct oxidation of Hg0 by lattice oxygen is energetically demanding within the present computational framework. Among the three sites, Hg–Ot gives the lowest activation energy and reaction energy, 146.817 and 134.941 kcal mol−1, respectively. Hg–Ob has the highest reaction energy, 163.136 kcal mol−1, while Hg–Ob′ gives the highest activation energy, 184.529 kcal mol−1. Among the three lattice-oxygen pathways examined, the Hg–Ot pathway has the lowest calculated activation barrier.
Figure 4.
Reaction energy profiles for Hg0 oxidation and oxidative desorption on V2O5/TiO2(001).
Figure 5.
Optimized reactant, transition-state, and product structures for Hg0 oxidation on V2O5/TiO2(001).
For the Hg–Ot pathway, Hg0 is first located above the terminal oxygen region, with an Hg–O distance of 3.524 Å. During the reaction, Hg gradually moves toward Ot, and the Hg–O distance decreases to about 2.771 Å at the transition state. In the final state, an HgO-like species forms, with an Hg–O bond length of about 2.042 Å. The pathway involves direct transfer of the terminal lattice O atom to Hg, followed by formation and detachment of an HgO-like product. A key factor behind the relatively low barrier is the structural degree of freedom of Ot: unlike the oxygen atoms locked at the bridging position, the terminal oxygen is less constrained by the surrounding skeleton, so its transfer from the surface requires less extensive structural rearrangement. The Hg–Ob reaction pathway follows a similar oxygen transfer process but entails a higher energy cost. In the reactant state, Hg0 is situated near the Ob atom of the V–O–V linkage; the Hg–O distance shortens from 3.222 Å in the reactant state to 2.600 Å in the transition state and further to 2.028 Å in the product state. Since the Ob atom acts as a structural bridge between two V centers, the removal of the oxygen atom disrupts the V–O–V framework and induces significant local structural distortion, consistent with the high calculated reaction energy. For the Hg–Ob′ pathway, the V–O–Ti environment at the interface undergoes more drastic restructuring. During progression toward the transition state, the terminal Ot atom migrates toward the Ob′ region while the local V–O–Ti framework rearranges. This coupled oxygen migration and interfacial restructuring gives the Hg–Ob′ pathway the highest calculated activation barrier among the three clean-surface pathways examined.
In general, on the V2O5/TiO2(001) surface, the direct oxidation of Hg0 initiated by lattice oxygen involves overcoming relatively high energy barriers for all three pathways investigated. Among them, the Hg–Ot pathway exhibits the lowest calculated activation barrier; in contrast, the Hg–Ob and Hg–Ob′ pathways entail higher barriers, which are attributed to the disruption of the V–O–V framework and the restructuring of the V–O–Ti interfacial environment, respectively.
3.2. Effect of NH3 on Hg0 Adsorption and Oxidation on V2O5/TiO2(001)
3.2.1. NH3 Adsorption on V2O5/TiO2(001)
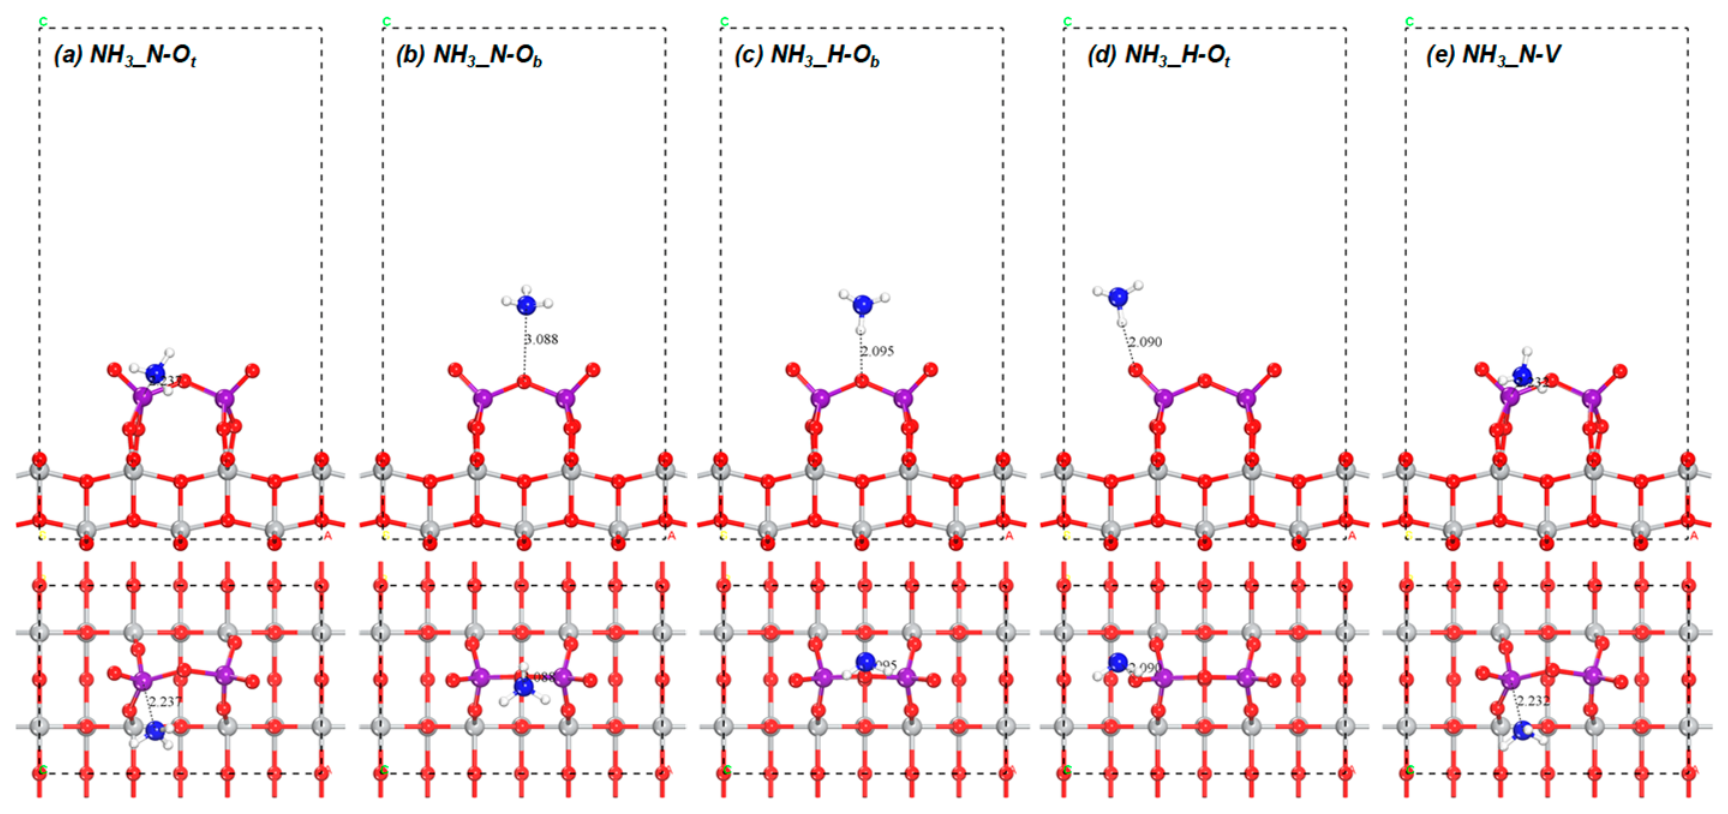
We studied the adsorption of NH3 on V2O5/TiO2 (001) to determine its optimal adsorption site and its interaction with surface vanadium–oxygen species. The adsorbed NH3 can occupy sites that Hg may also utilize and can change the local electronic environment for the later adsorption and oxidation of Hg. Five initial configurations are considered: NH3_N-Ot, NH3_N-Ob, NH3_H-Ob, NH3_H-Ot, and NH3_N-V, as shown in Figure 6. Their adsorption energies, main structural parameters and Mulliken charges are listed in Table 3. These models compare two adsorption modes: N-directed adsorption (N lone pair electrons interacting with the surface) and H-directed adsorption (the interaction between H atoms and surface oxygen species).
Figure 6.
Optimized configurations of NH3 adsorbed on V2O5/TiO2(001).
Table 3.
Adsorption energies, characteristic distances, and Mulliken charges for NH3 adsorption on V2O5/TiO2(001).
The N-oriented model with NH3 close to the exposed vanadium center is obviously the most stable. The NH3_N-Ot and NH3_N-V configurations exhibit more negative adsorption energies of −32.139 and −32.285 kcal mol−1, respectively, with characteristic adsorption distances of 2.237 and 2.232 Å and Mulliken charges of 0.324 and 0.328 e, respectively. Although the two initial configurations are different, the structural relaxation drives them toward essentially the same final geometry, indicating that the V-Ot local environment is essentially suitable for strong NH3 coordination. In contrast, the interaction between ammonia nitrogen and bridge oxygen (NH3_N-Ob) is much weaker: the adsorption energy is reduced to only −10.452 kcal mol−1, the optimized distance is extended to 3.088 Å, and the charge transfer amount is moderate at 0.126 e. Unlike the exposed vanadium sites, the bridging oxygen in the V-O-V bond does not act as an accessible electron-pair acceptor (a Lewis acid center) and cannot strongly bind to the nitrogen lone pair electrons. When ammonia interacts with surface oxygen through its hydrogen atom, the bonding strength is between these extremes, but remains moderate. The adsorption energies of NH3_H-Ob and NH3_H-Ot are −14.118 and −13.692 kcal mol−1, respectively. There is a similar short contact distance near 2.09 Å, and the charges are 0.163 e and 0.146 e, respectively. These values are significantly lower, i.e., substantially less negative, than those of the stable N-coordination configuration, and the nature of the interaction is better described as similar to a stable hydrogen bond, rather than a strong donor–acceptor bond.
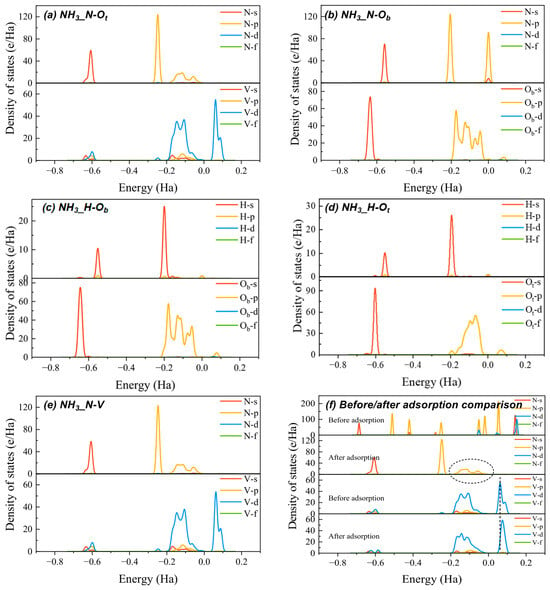
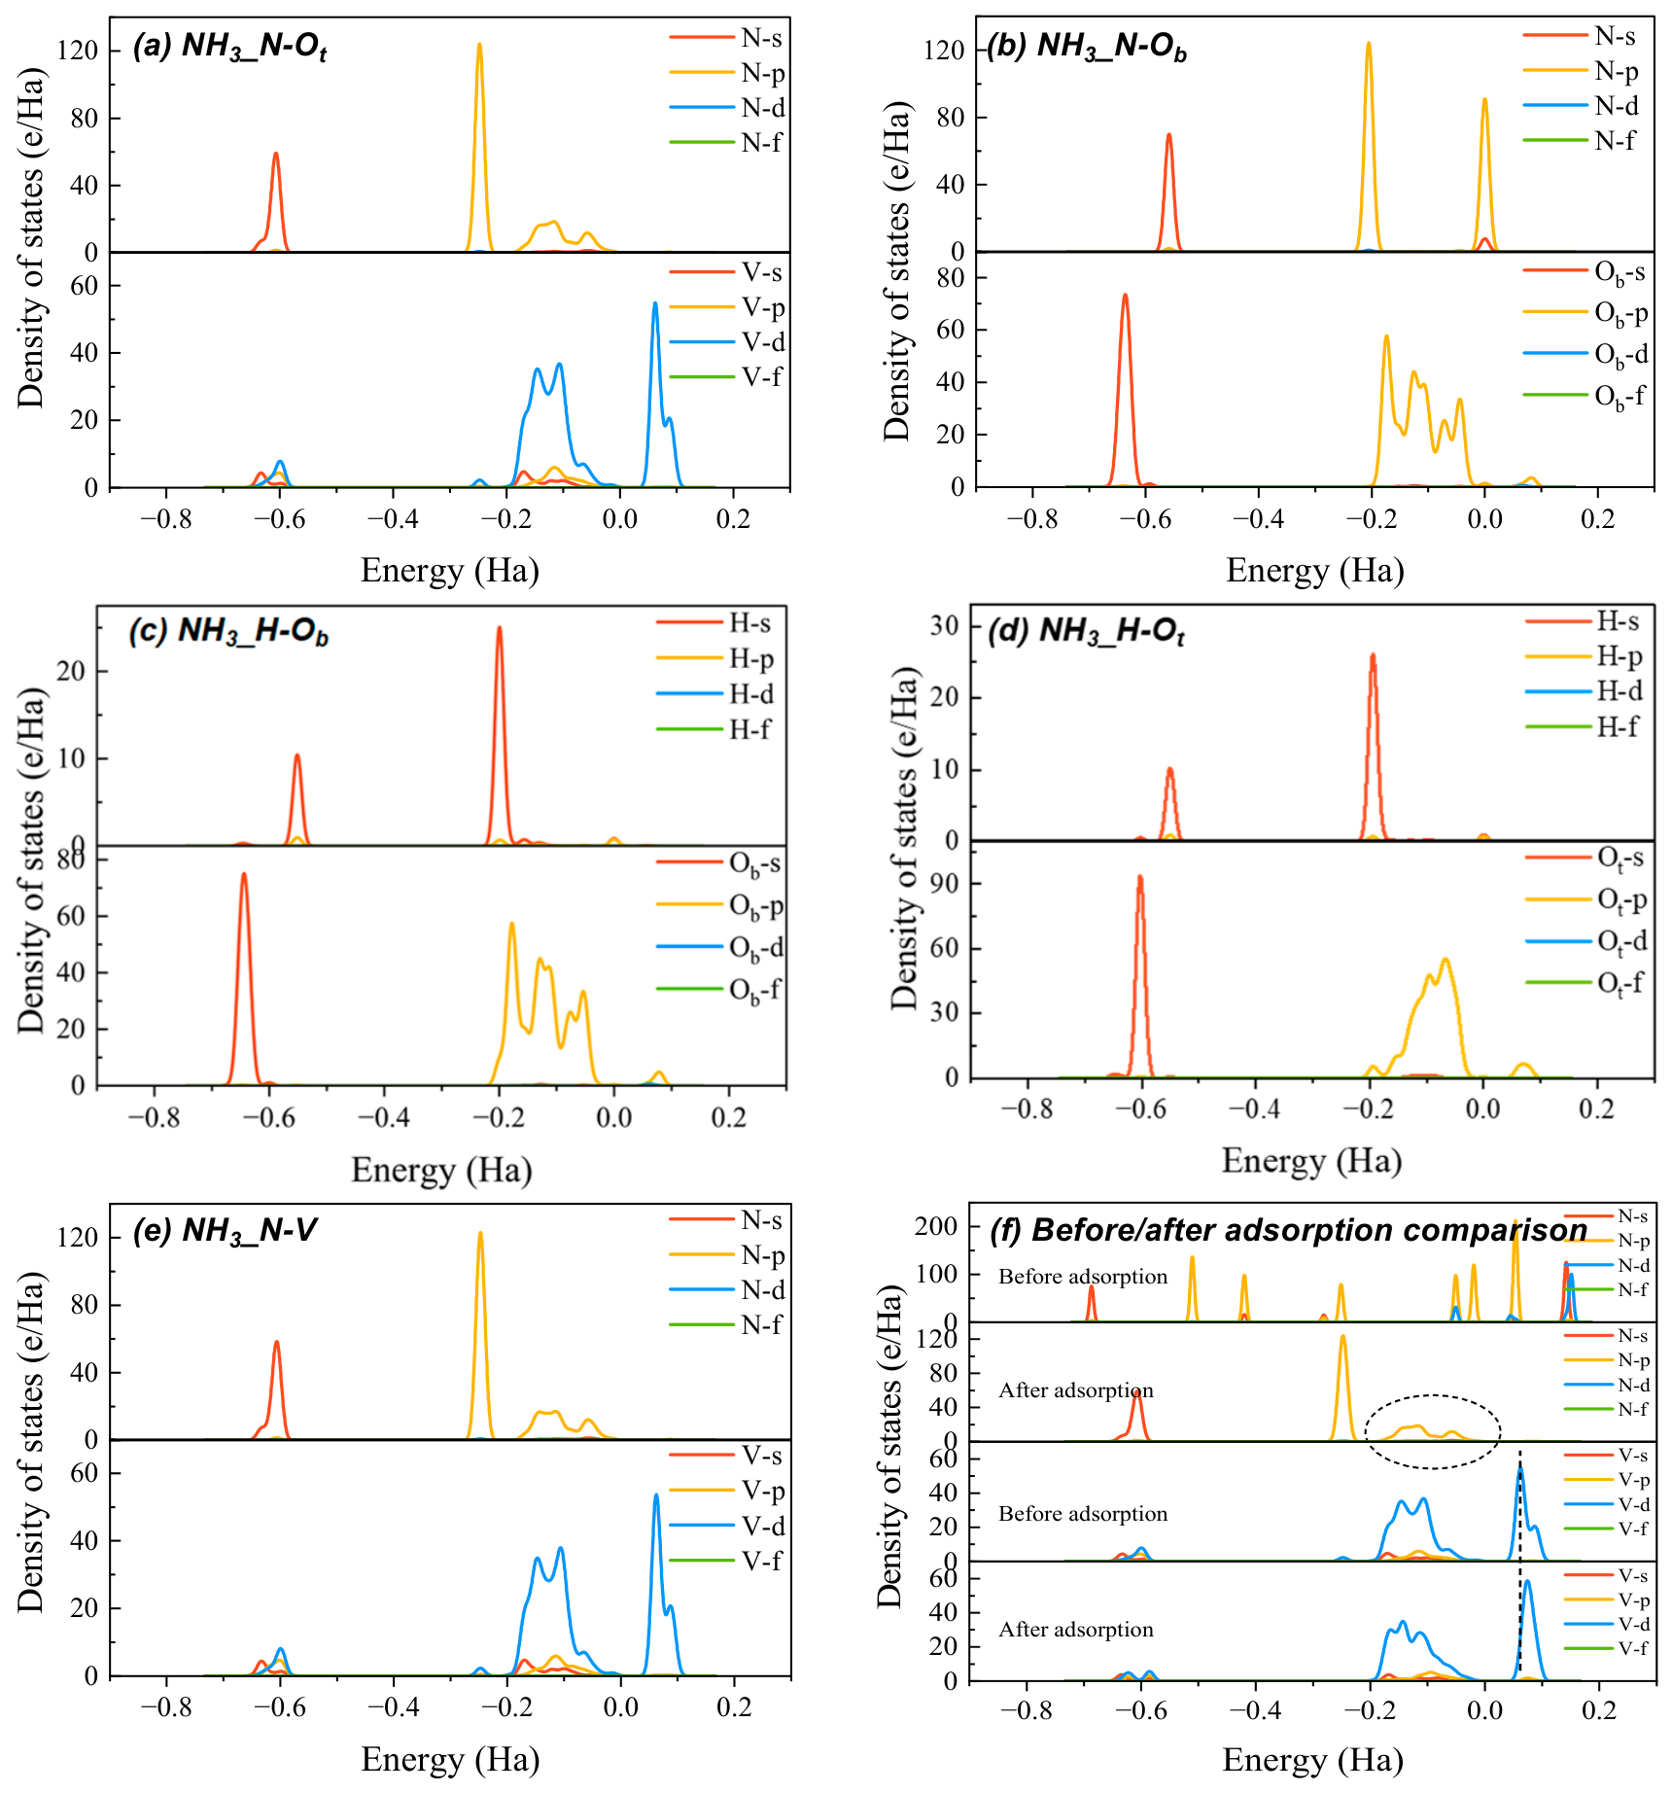
The PDOS results in Figure 7 explain the electronic origin of this adsorption trend. As shown in Figure 7a–e, clear adsorbate–surface hybridization appears only in the NH3_N–Ot and NH3_N–V configurations, whereas the other three cases show no obvious hybridization features. Since NH3_N–Ot and NH3_N–V relax to nearly the same stable adsorption structure, we selected NH3_N–V as the representative case for comparing the PDOS before and after adsorption, as shown in Figure 7f, to represent the stable N-coordinated state. After NH3 adsorption, a new feature appears in the N p states, accompanied by obvious hybridization with V d states. Meanwhile, the V d states shift after adsorption, demonstrating that the exposed V center is electronically perturbed by NH3 coordination. These PDOS results are fully consistent with the adsorption energies, optimized distances, and Mulliken charge analysis. NH3 adsorption at NH3_N–V is mainly governed by direct N(p)–V(d) interaction between the NH3 lone pair and the exposed V center, rather than by simple electrostatic attraction. This orbital interaction accounts for the much higher stability of NH3_N–V and NH3_N–Ot compared with the Ob-centered and H-oriented configurations. Overall, NH3 preferentially adsorbs at exposed V-related sites, especially in the V–Ot local environment, whereas adsorption near the Ob region or through H-oriented interaction is relatively weak.
Figure 7.
PDOS analysis of NH3 adsorption on V2O5/TiO2(001).
3.2.2. Effect of Pre-Adsorbed NH3 on Hg0 Adsorption and Oxidation
Hg0 Adsorption Behavior
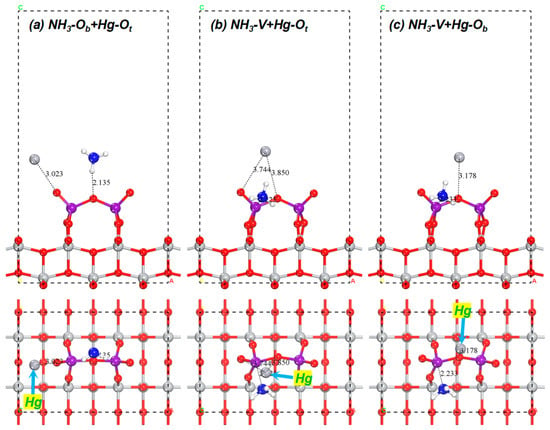
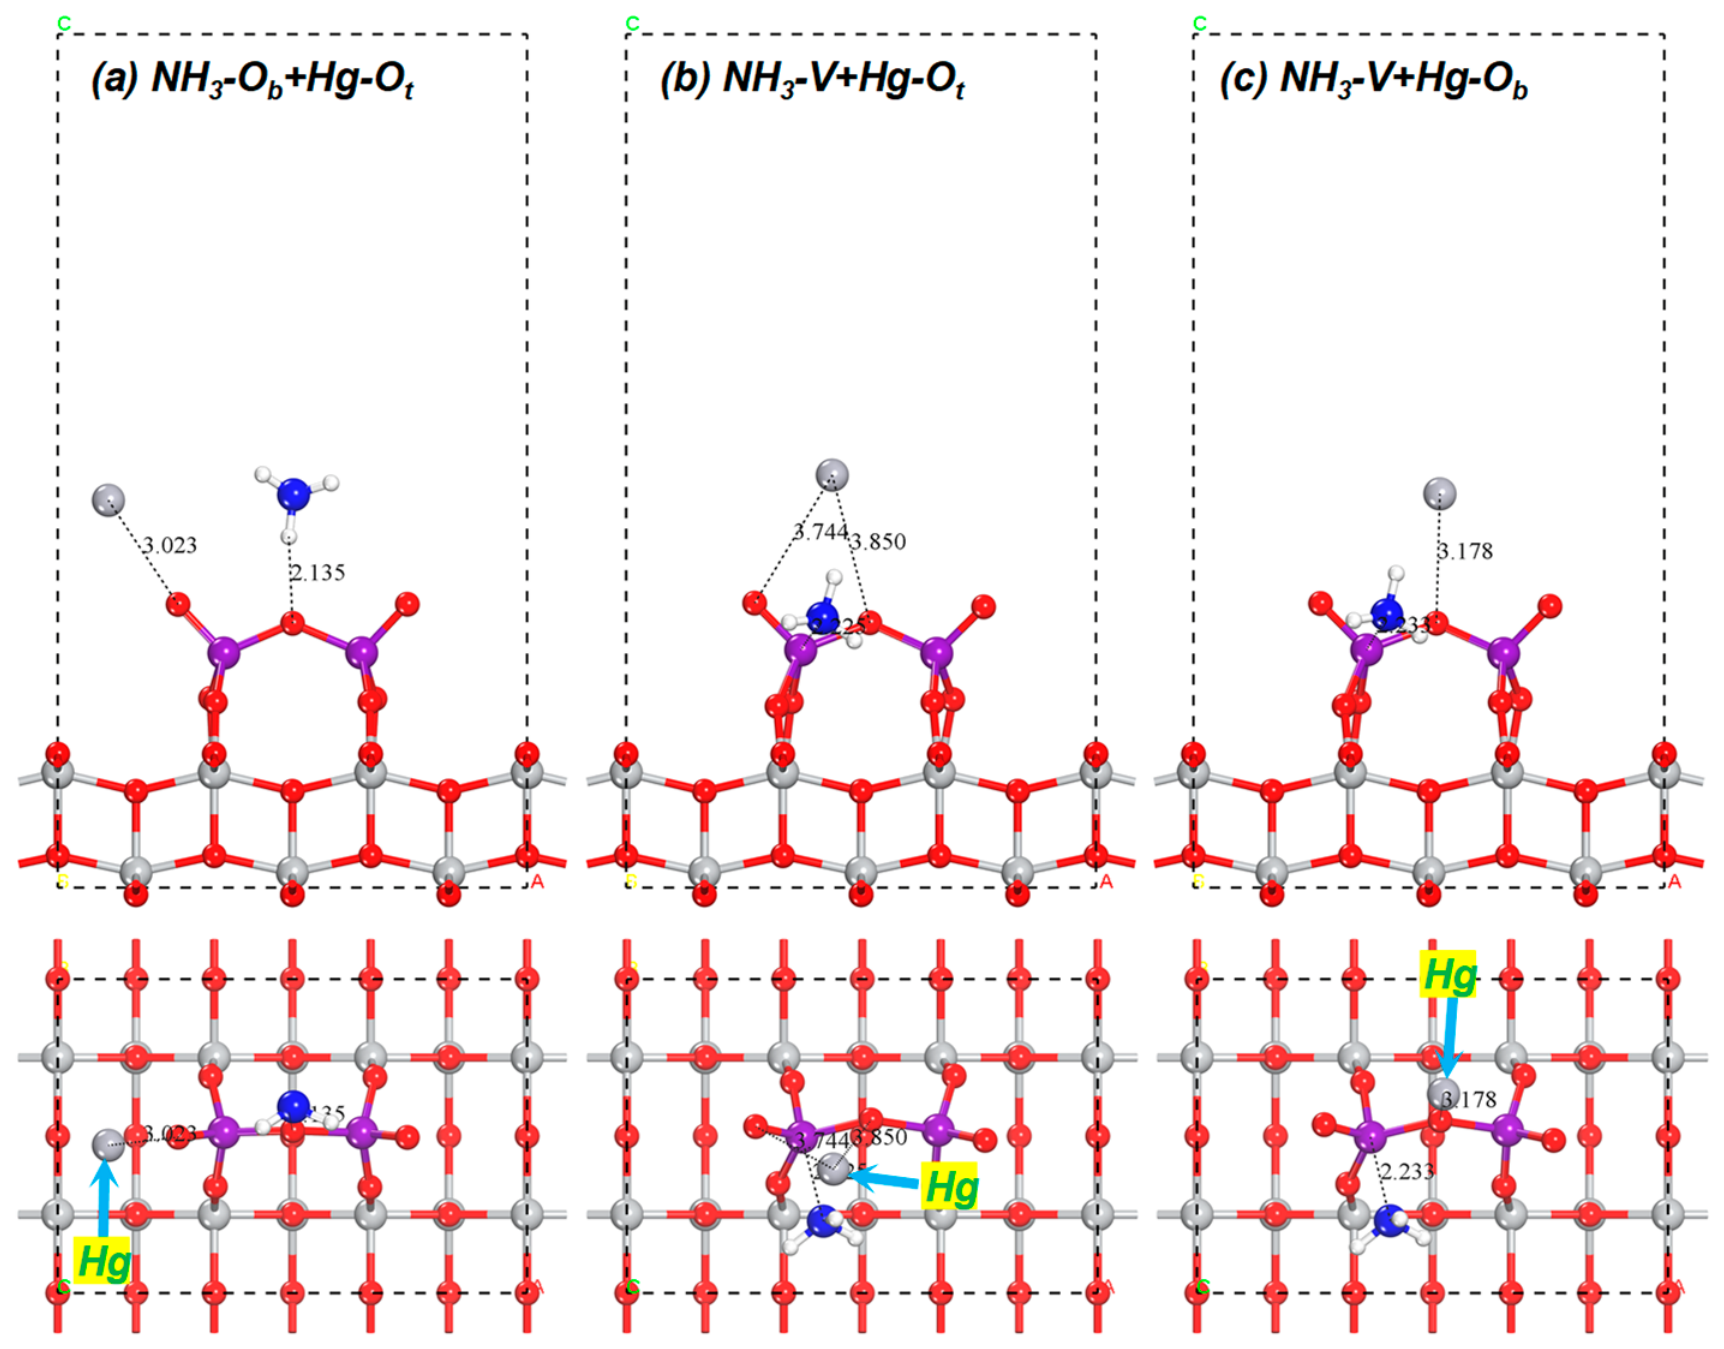
The effect of pre-adsorbed NH3 on Hg0 adsorption was examined using three representative co-adsorption configurations on V2O5/TiO2(001), as shown in Figure 8. In the notation used here, NH3–Ob+ Hg–Ot denotes a configuration in which NH3 is first adsorbed in the Ob-related region, followed by Hg0 adsorption near the Ot site. The same notation is used for the other co-adsorption models. Two types of NH3-precovered surfaces were considered. The first involves NH3 occupation of an oxygen-related region, represented by NH3–Ob, and mainly reflects local site blocking. The second involves N-coordinated NH3 adsorption at an exposed V-related site, represented by NH3_N–V, where both the coordination structure and the electronic environment of neighboring V–O units may be modified. The corresponding adsorption energies, characteristic distances, and Mulliken charges are listed in Table 4.
Figure 8.
Optimized co-adsorption configurations of Hg0 on NH3-precovered V2O5/TiO2(001).
Table 4.
Adsorption energies, characteristic distances, and Mulliken charges for Hg0 adsorption on NH3-precovered V2O5/TiO2(001).
Compared with the clean surface, the adsorption of Hg0 is weakened in all NH3 pre-adsorbed configurations. The adsorption energies of Hg-Ot and Hg-Ob on the intrinsic V2O5/TiO2(001) surface are −12.31 kcal mol−1 and −12.58 kcal mol−1, respectively. These adsorption energies become less negative after NH3 pre-adsorption, reaching −11.04 (NH3-Ob + Hg-Ot), −11.50 kcal mol−1 (NH3_N-V + Hg-Ot) and −11.84 kcal mol−1 (NH3_N-V + Hg-Ob). A similar trend is observed in the Mulliken charge analysis. The charge on Hg0 decreases from 0.113 e and 0.115 e on the clean Hg-Ot and Hg-Ob surfaces to 0.088 e, 0.053 e and 0.056 e in the three NH3 precovered configurations. The less negative adsorption energies and reduced charge transfer indicate that the pre-adsorbed NH3 weakens the interaction between Hg0 and V2O5/TiO2(001) surfaces.
For the NH3-Ob+ Hg-Ot configuration, the optimized Hg0-surface distance is 3.023 Å, and the adsorption energy is −11.04 kcal mol−1. Although Hg0 is still close to the Ot region, NH3 located in the adjacent Ob-related region weakens its adsorption stability. This behavior is mainly related to the local occupation of the adsorption environment, which limits the entry of Hg0 into the surface area near the potential adsorption site. This effect becomes more pronounced when NH3 is pre-adsorbed in the V-related region. For the NH3_N-V + Hg-Ot configuration, the optimized Hg0-surface distance increases to 3.744 and 3.850 Å, the adsorption energy decreases to −11.50 kcal mol−1, and the Mulliken charge on Hg0 decreases to 0.053 e. In the NH3_N-V + Hg-Ob configuration, the adsorption energy is −11.84 kcal mol−1, and the Mulliken charge on Hg0 is also small, at 0.056 e. Compared with the clean surface, the larger Hg0-surface distance and the lower charge transfer value indicate that the stability of Hg0 is weakened after NH3 coordination, especially at the exposed V-related sites. Therefore, once NH3 occupies the local environment around the V center, Hg0 is unable to interact as effectively with the adjacent V–O structure as it does on the clean surface.
Overall, NH3 pre-adsorption weakens Hg0 adsorption through two related effects. The first is competitive site occupation, in which NH3 partially blocks surface regions involved in Hg0 adsorption. The second is local electronic regulation, which becomes more pronounced when NH3 coordinates directly to an exposed vanadium site. In this case, the reduced adsorption energies and smaller Hg charge transfer values reflect a less favorable local coordination and electronic environment for Hg0 stabilization. Therefore, NH3 mainly suppresses Hg0 conversion at the initial adsorption stage by weakening the formation of stable Hg0 adsorption precursors. This finding provides a possible atomic-scale interpretation of the inhibitory effect of NH3 on Hg0 oxidation previously observed over vanadium-based SCR catalysts [8,12].
Hg0 Oxidative Desorption Behavior
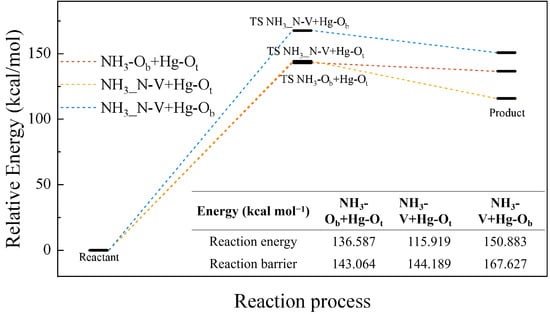
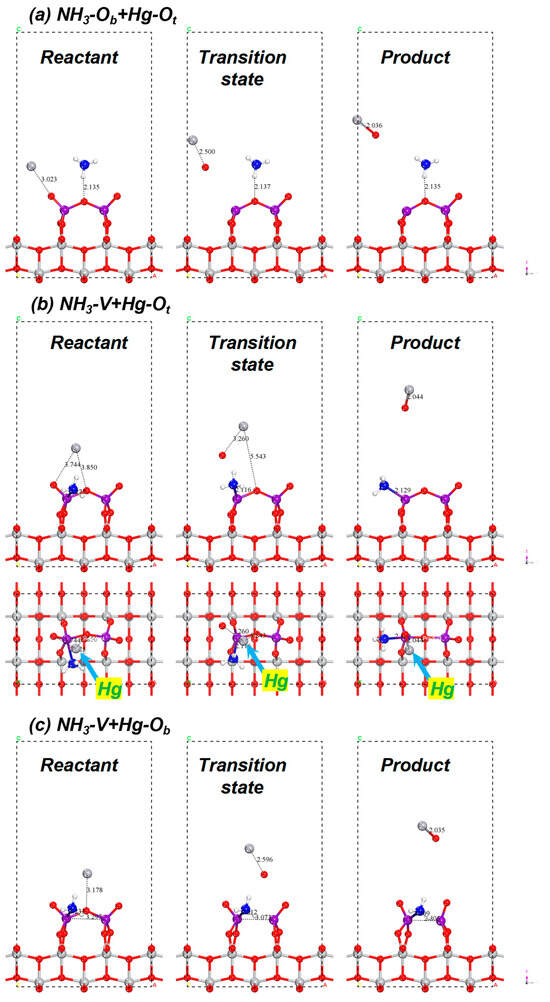
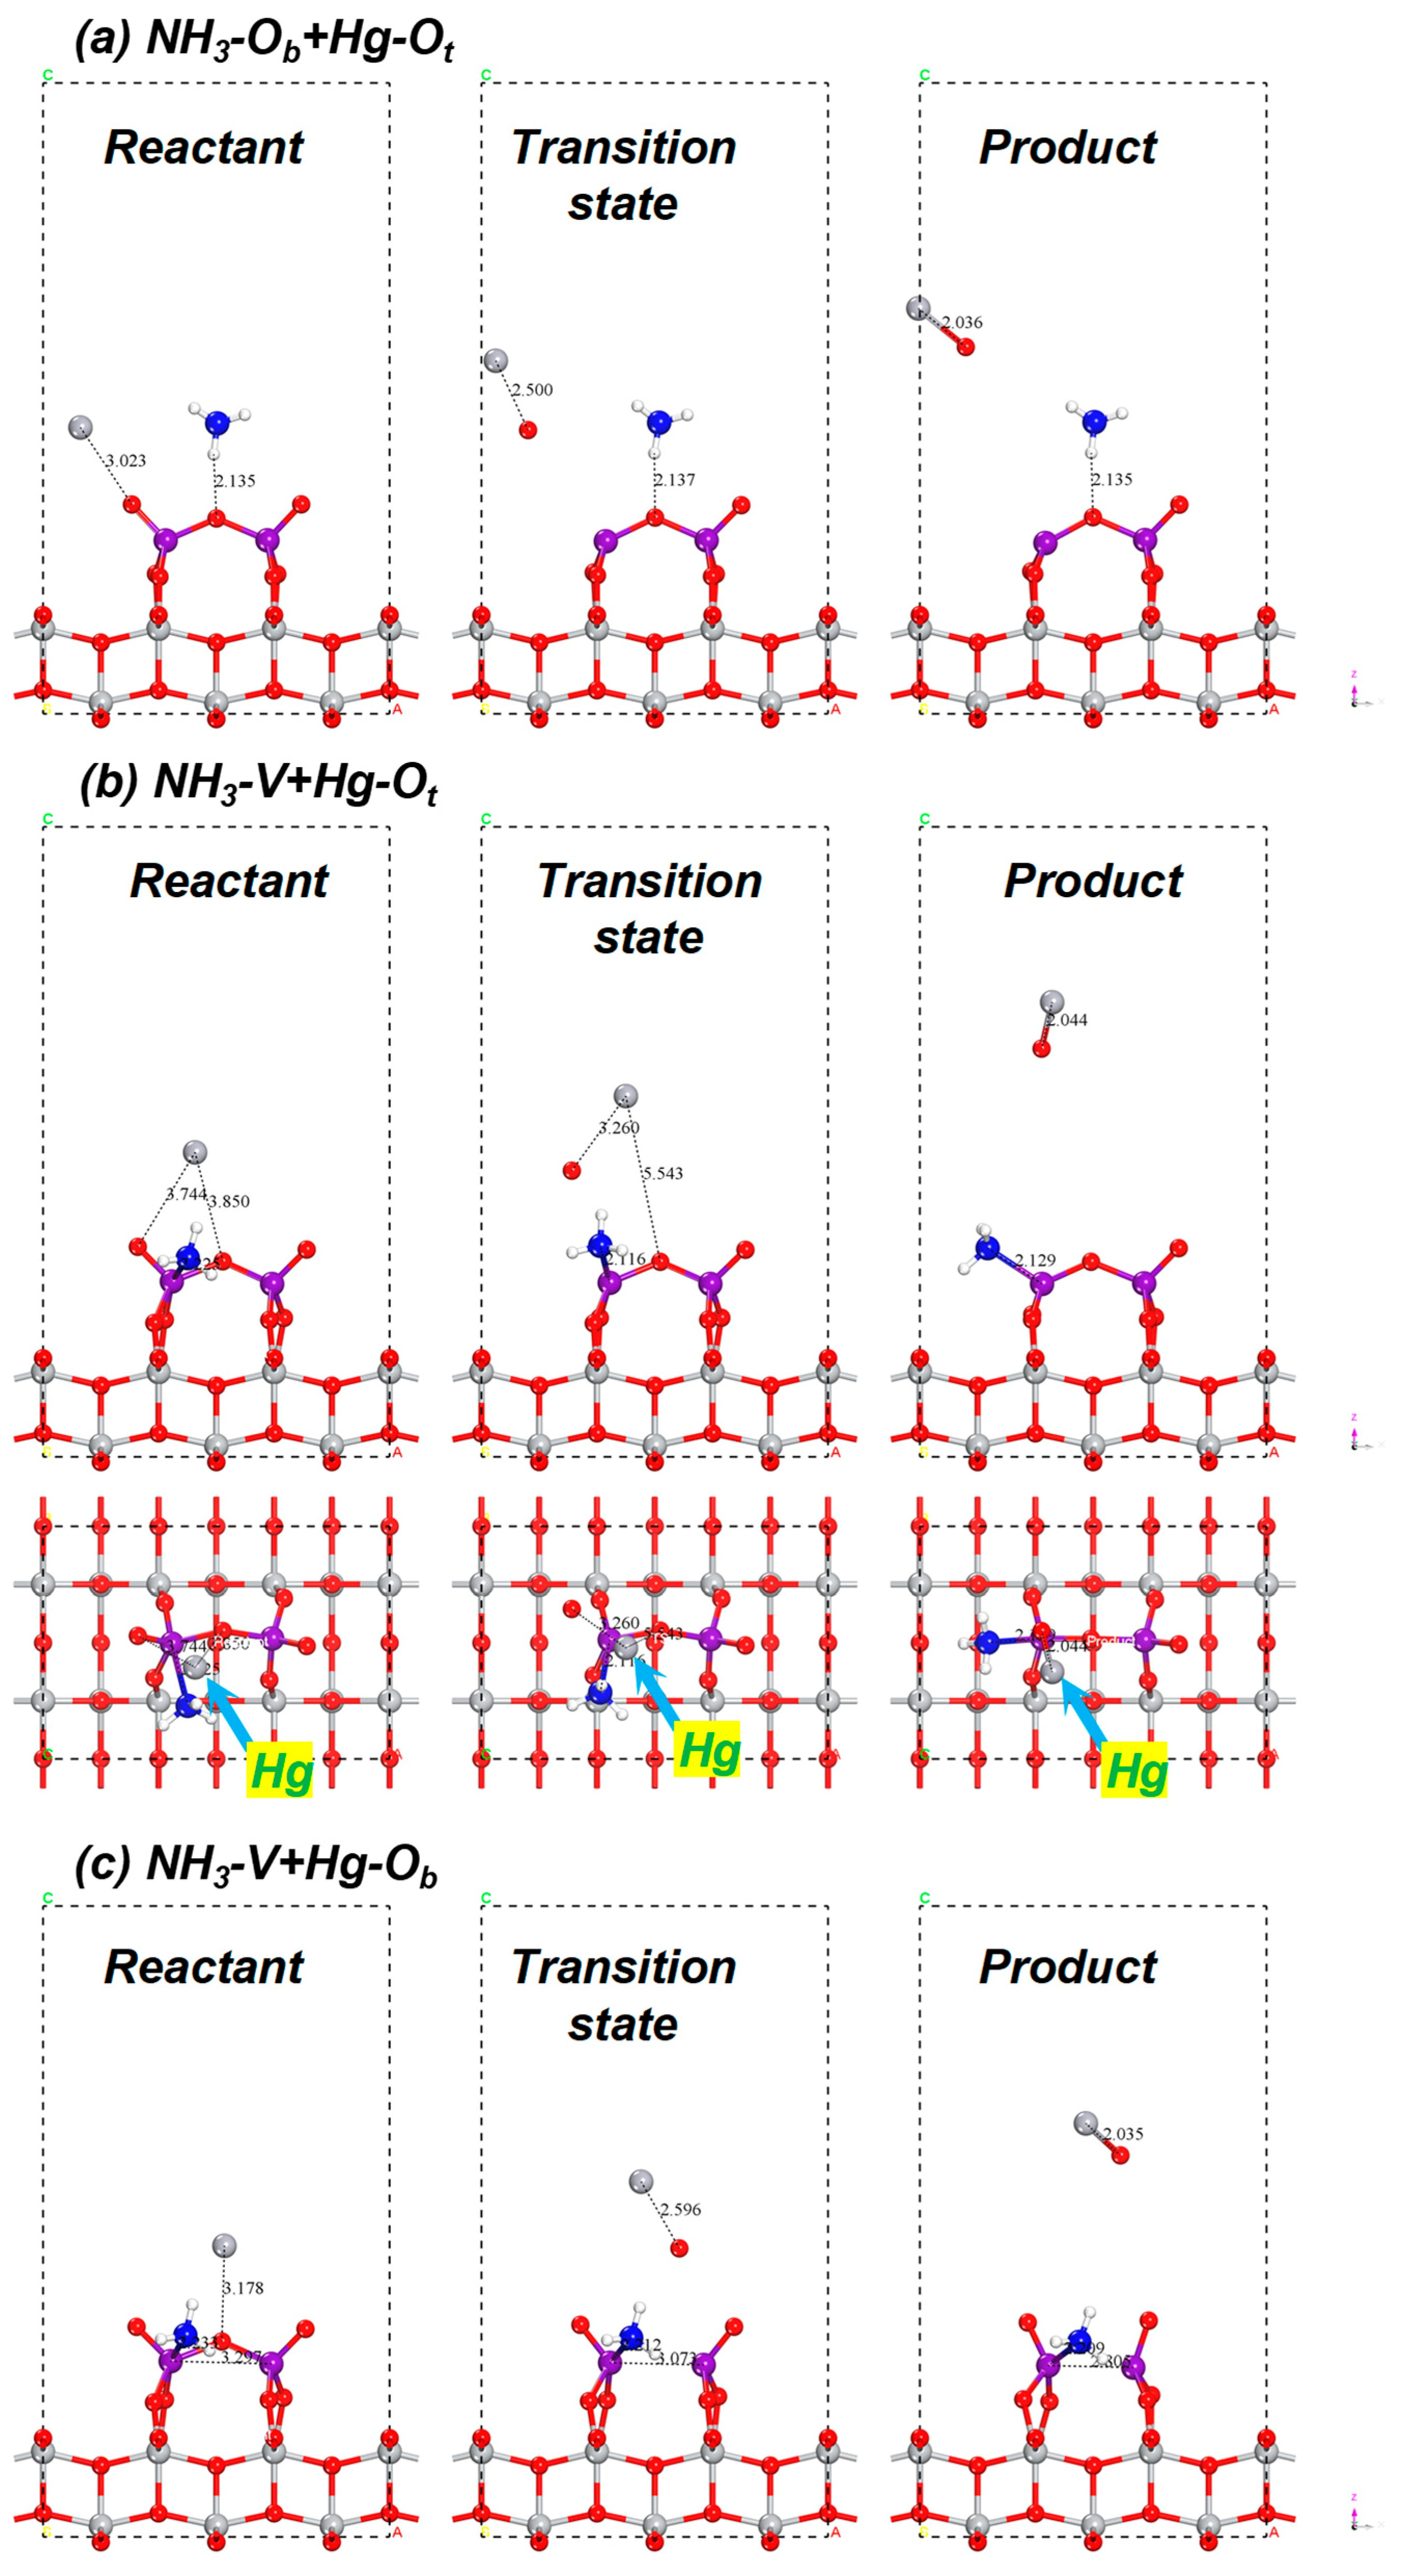
The subsequent oxidation and oxidative desorption of Hg on NH3-precovered V2O5/TiO2(001) surfaces were further examined by transition-state calculations. The corresponding reaction energy profiles are shown in Figure 9, and the optimized structures along the reaction pathways are presented in Figure 10. Three representative pathways were considered, namely NH3–Ob + Hg–Ot, NH3_N–V + Hg–Ot, and NH3_N–V + Hg–Ob. Similar to the clean-surface routes, all three pathways are highly endothermic and require large activation barriers. Therefore, Hg0 oxidation on the NH3-precovered surface remains energetically demanding within the present computational framework, while the overall oxygen-transfer/product-detachment mechanism remains similar to that on the clean surface.
Figure 9.
Reaction energy profiles for Hg0 oxidation on NH3-precovered V2O5/TiO2(001).
Figure 10.
Optimized reactant, transition-state, and product structures for Hg0 oxidation on NH3-precovered V2O5/TiO2(001).
For the NH3-Ob + Hg-Ot pathway, Hg0 is initially located above the terminal oxygen region, while NH3 is pre-adsorbed at the adjacent Ob site. During the reaction, Hg moves to the active oxygen, and the Hg-O distance decreases from 3.023 Å to about 2.500 Å at the transition state. In the final state, a HgO-like species with a Hg-O bond length of about 2.036 Å is formed, while NH3 remains near the V-O unit. The reaction energy and activation energy are 136.587 and 143.064 kcal mol−1, respectively, which are close to the values of the clean Hg-Ot pathway. This indicates that the NH3 at the Ob site does not block the oxygen transfer from Ot to Hg, but only slightly changes the energy of the reaction. The NH3_N-V + Hg-Ot pathway emphasizes the role of NH3 coordination at the exposed V site. Here, NH3 binds to the V center, and Hg0 begins near the Ot region. As the reaction proceeds, Hg approaches the terminal oxygen, and the Hg-O distance decreases from 3.744/3.850 Å to about 2.269 Å at the transition state. The final structure forms a HgO-like product with a Hg- O bond length of about 2.044 Å. The reaction energy and activation energy are 115.919 and 144.189 kcal mol−1, respectively. The activation barrier is similar to that on the clean surface, but the reaction energy is lower. Following the formation and detachment of the HgO-like product, NH3 shifts from the original position, indicating that the local V-related environment relaxes after the reaction. The NH3_N-V + Hg-Ob pathway follows similar oxidation and desorption processes but is not as favorable as the Ot-related pathway. The Hg-O distance decreases from 3.178 Å to about 2.596 Å in the transition state. In the final state, the Hg-O bond length is about 2.035 Å. The reaction energy and activation energy are 150.883 and 167.627 kcal mol−1, respectively. These values are lower than those of the clean Hg-Ob pathway, indicating that V-site NH3 coordination can partially reduce the energy cost. However, the bridging oxygen pathway still exhibits a comparatively high calculated barrier because substantial distortion of the V–O–V framework is required.
Combining the transition state and adsorption analysis results, NH3 primarily influences the Hg0 oxidation conversion during the initial adsorption stage. The pre-adsorbed NH3 weakens the ability of Hg to be captured, as evidenced by the smaller negative adsorption energy and the reduced Hg charge transfer. Once Hg0 is already near the reactive oxygen site, the subsequent oxygen-transfer/product-detachment process is similar to that on the clean surface.
3.3. Effect of NO2 on Hg0 Adsorption and Oxidation over V2O5/TiO2(001)
3.3.1. NO2 Adsorption on V2O5/TiO2(001)
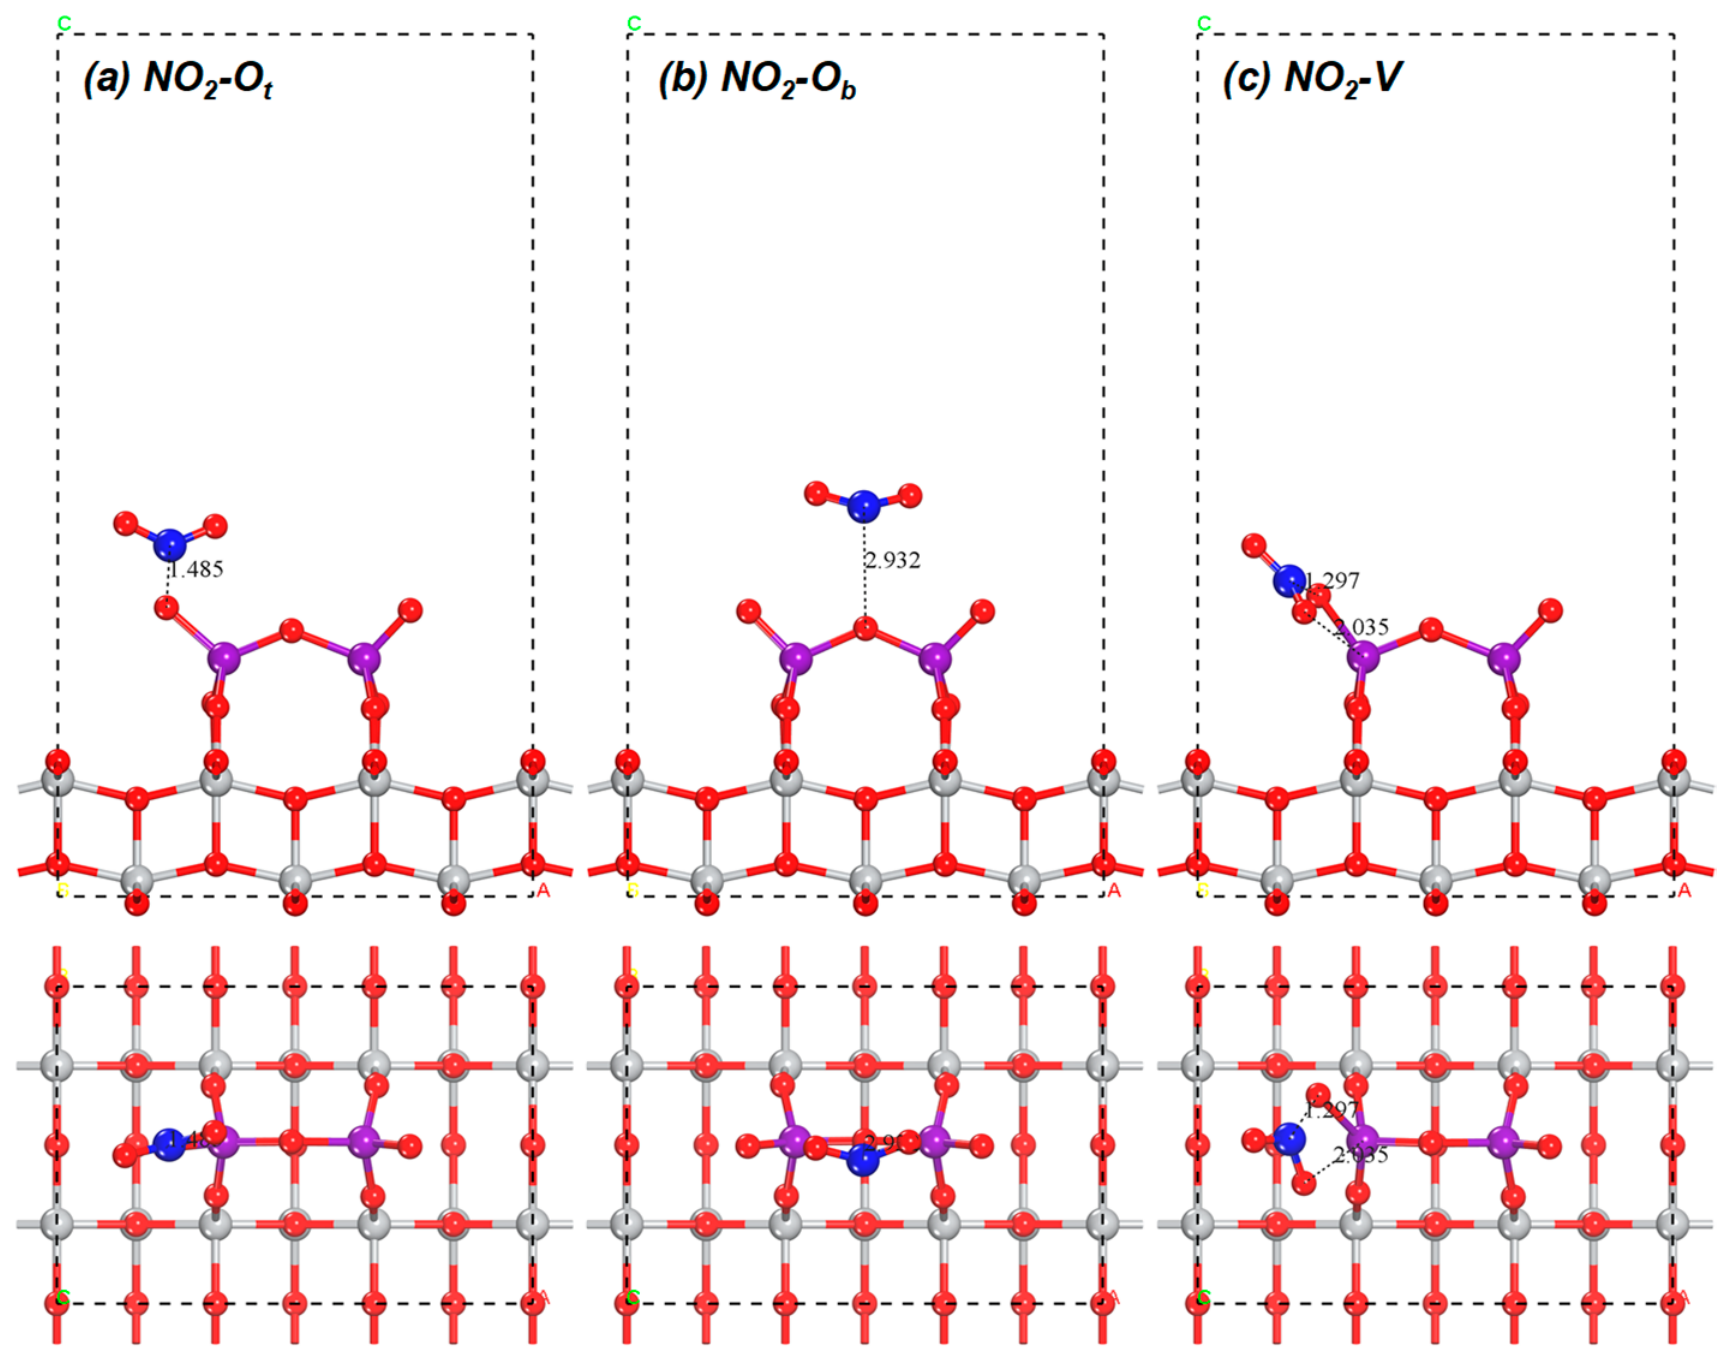
We studied the adsorption of NO2 on V2O5/TiO2(001) to identify its preferred adsorption configuration and its effect on the local V–O environment. As a key component in the FSCR atmosphere, NO2 may not only participate in gas-phase reactions but also alter the redox and electronic properties of vanadium-based catalysts. Its effect depends strongly on its adsorption geometry, because different geometries disturb different V–O units and change the local distribution of reactive oxygen species. Three representative initial configurations were considered: NO2-Ot, NO2-Ob and NO2-V, as shown in Figure 11. Their adsorption energies, characteristic distances and Mulliken charges are summarized in Table 5.
Figure 11.
Optimized configurations of NO2 adsorbed on V2O5/TiO2(001).
Table 5.
Adsorption energies, characteristic distances, and Mulliken charges for NO2 adsorption on V2O5/TiO2(001).
The results of adsorption energy show that NO2 is more inclined to adsorb in the exposed V-related region. Among the three configurations, NO2-V has the most negative adsorption energy, at −10.028 kcal mol−1, followed by NO2-Ob, which has an adsorption energy of −7.938 kcal mol−1. The stability of NO2-Ot is poor, and its adsorption energy is only −2.969 kcal mol−1. Optimizing the structure supports this trend. In NO2-Ot, the characteristic distance is 1.485 Å, indicating that NO2 is in close contact with the terminal oxygen region. In NO2-Ob, NO2 is located above the bridging oxygen region with a characteristic distance of 2.932 Å. In NO2-V, two characteristic distances are observed: O(NO2)-V_surface is 2.035 Å, and N(NO2)-Ot_surface is 1.297 Å. This means that the relaxed NO2-V structure interacts with the exposed V center and the adjacent terminal oxygen at the same time, which explains its higher stability.
Mulliken charge analysis shows that NO2 adsorption involves limited charge transfer. The calculated charges of NO2-Ot, NO2-Ob and NO2-V adsorbing NO2 are 0.126 e, 0.081 e and 0.085 e, respectively. Although charge transfer occurs in all three cases, the magnitude is still very small. The most stable NO2-V configuration does not exhibit the maximum charge transfer. This indicates that the adsorption strength is not solely controlled by net electron transfer. On the contrary, the local geometric and cooperative interactions between NO2, the V center and nearby oxygen atoms play a major role in stabilizing NO2 adsorption.
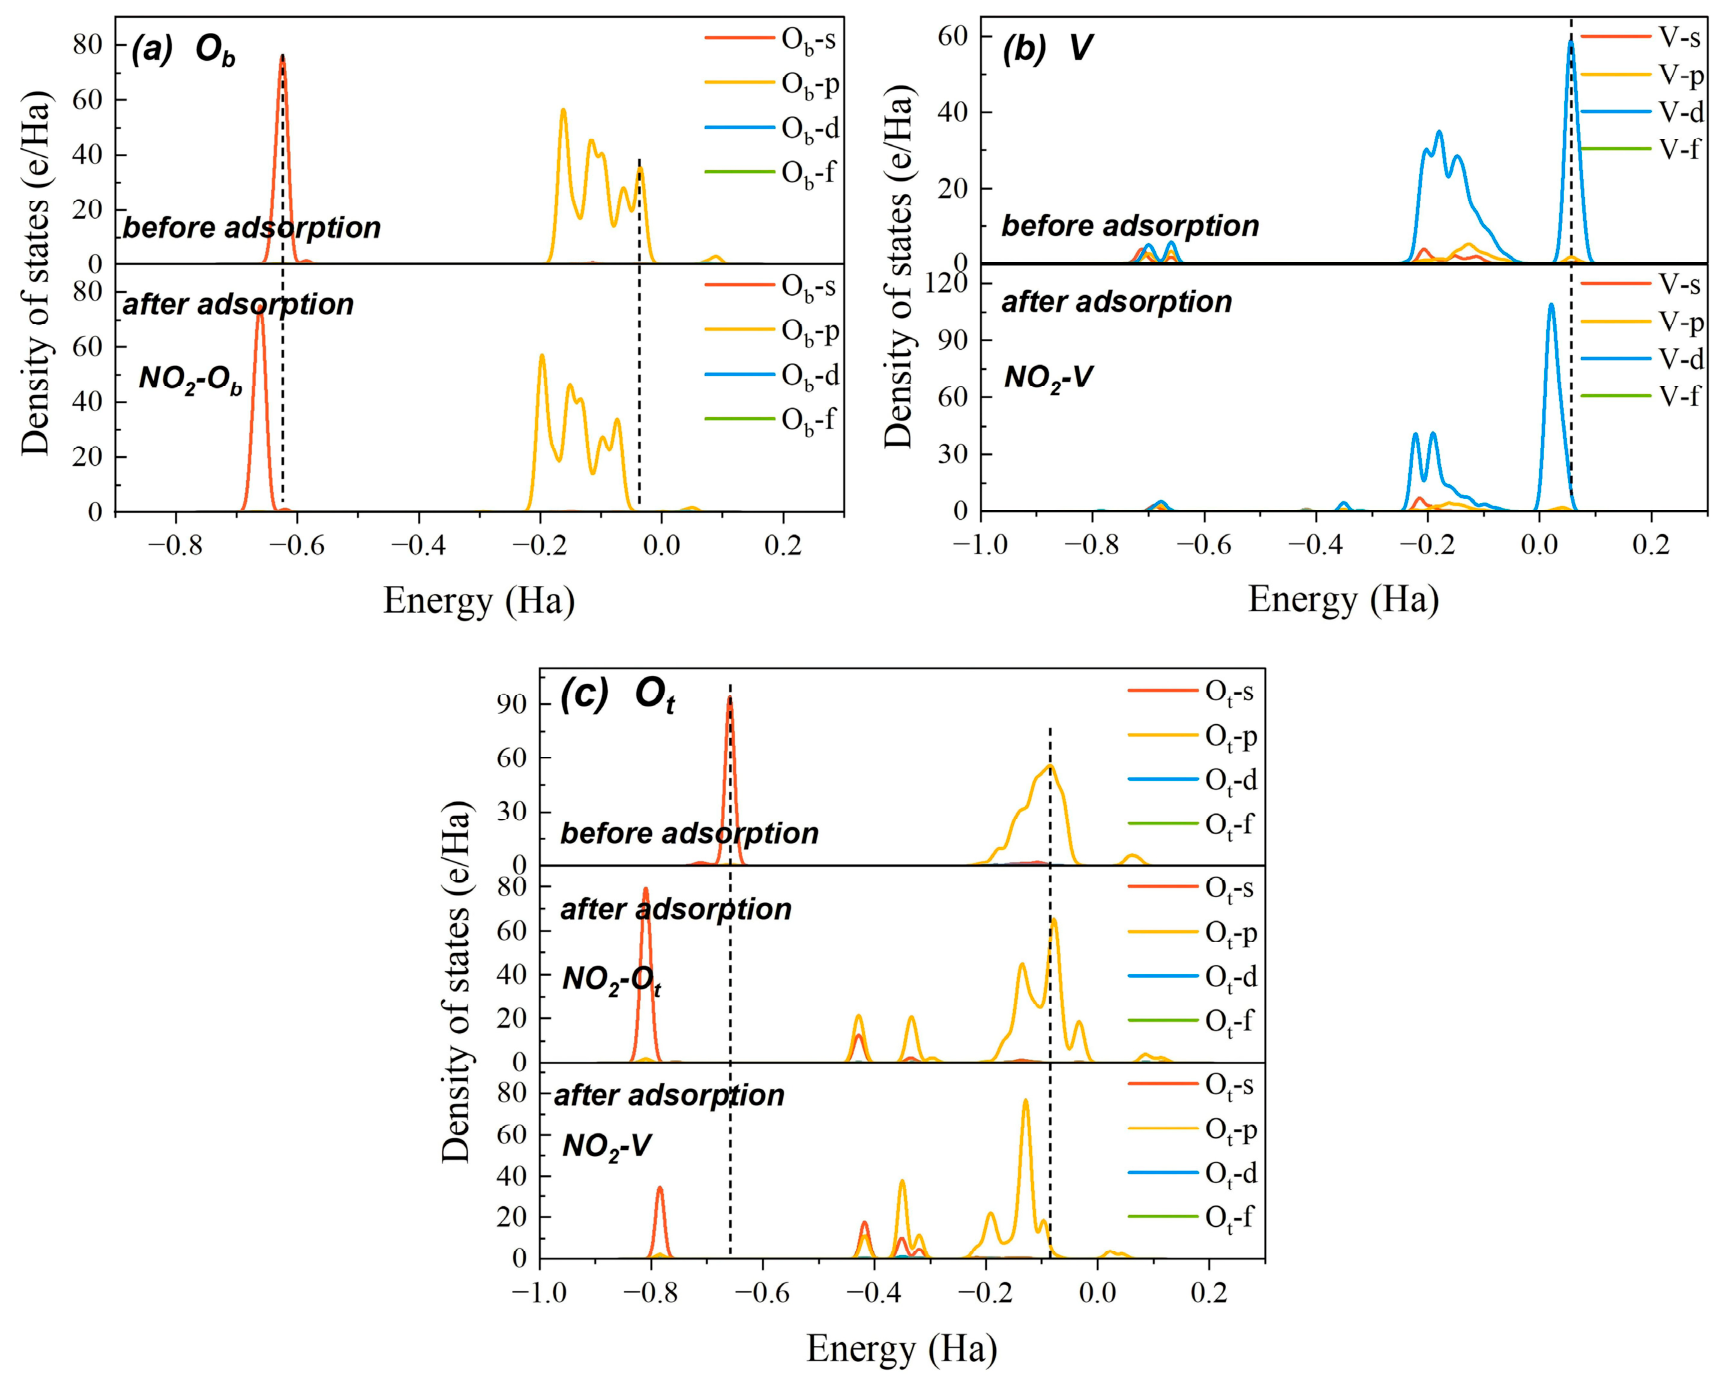
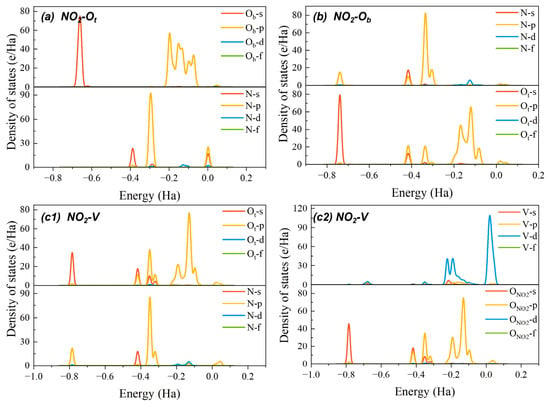
The PDOS results further clarify the electron source of NO2 stability. As shown in Figure 12, the adsorption of NO2 causes a significant change in the local electronic structure of the catalyst surface. In NO2-Ob, the PDOS of Ob atoms changes significantly after adsorption, especially the peak intensity and energy distribution, which manifests as an electronic disturbance in the V-O-V bridging region. In NO2-V, the V-related states change more strongly, especially the V d state, which indicates that there is a direct electronic interaction between NO2 and the exposed V site. After the adsorption of NO2-Ot and NO2-V, the Ot-related states also change. This indicates that the terminal oxygen environment can be affected both by the direct contact of NO2 and the adsorption of NO2 near the V center. The atomic and orbital-resolved PDOS in Figure 13 identify the orbital coupling responsible for NO2 stabilization. For NO2–Ot, the N p states of NO2 overlap with the p states of the nearby surface Ot atoms, demonstrating local electronic coupling between NO2 and the terminal oxygen region. In NO2–Ob, the N p states of NO2 overlap with the p states of the Ob surface, showing a similar coupling. The strongest orbital interactions occur in the NO2–V configuration. In this configuration, the N-related states of NO2 couple with the adjacent Ot-related states, while the O p states of NO2 show more pronounced overlap with the V d states on the catalyst surface. This synergistic N(NO2)–Ot and O(NO2)–V interaction explains the higher stability of NO2–V.
Figure 12.
PDOS comparison of selected surface atoms before and after NO2 adsorption on V2O5/TiO2(001).
Figure 13.
Atom- and orbital-resolved PDOS of NO2 adsorption systems on V2O5/TiO2(001).
3.3.2. Effect of Pre-Adsorbed NO2 on Hg0 Adsorption and Oxidation
Hg0 Adsorption Behavior
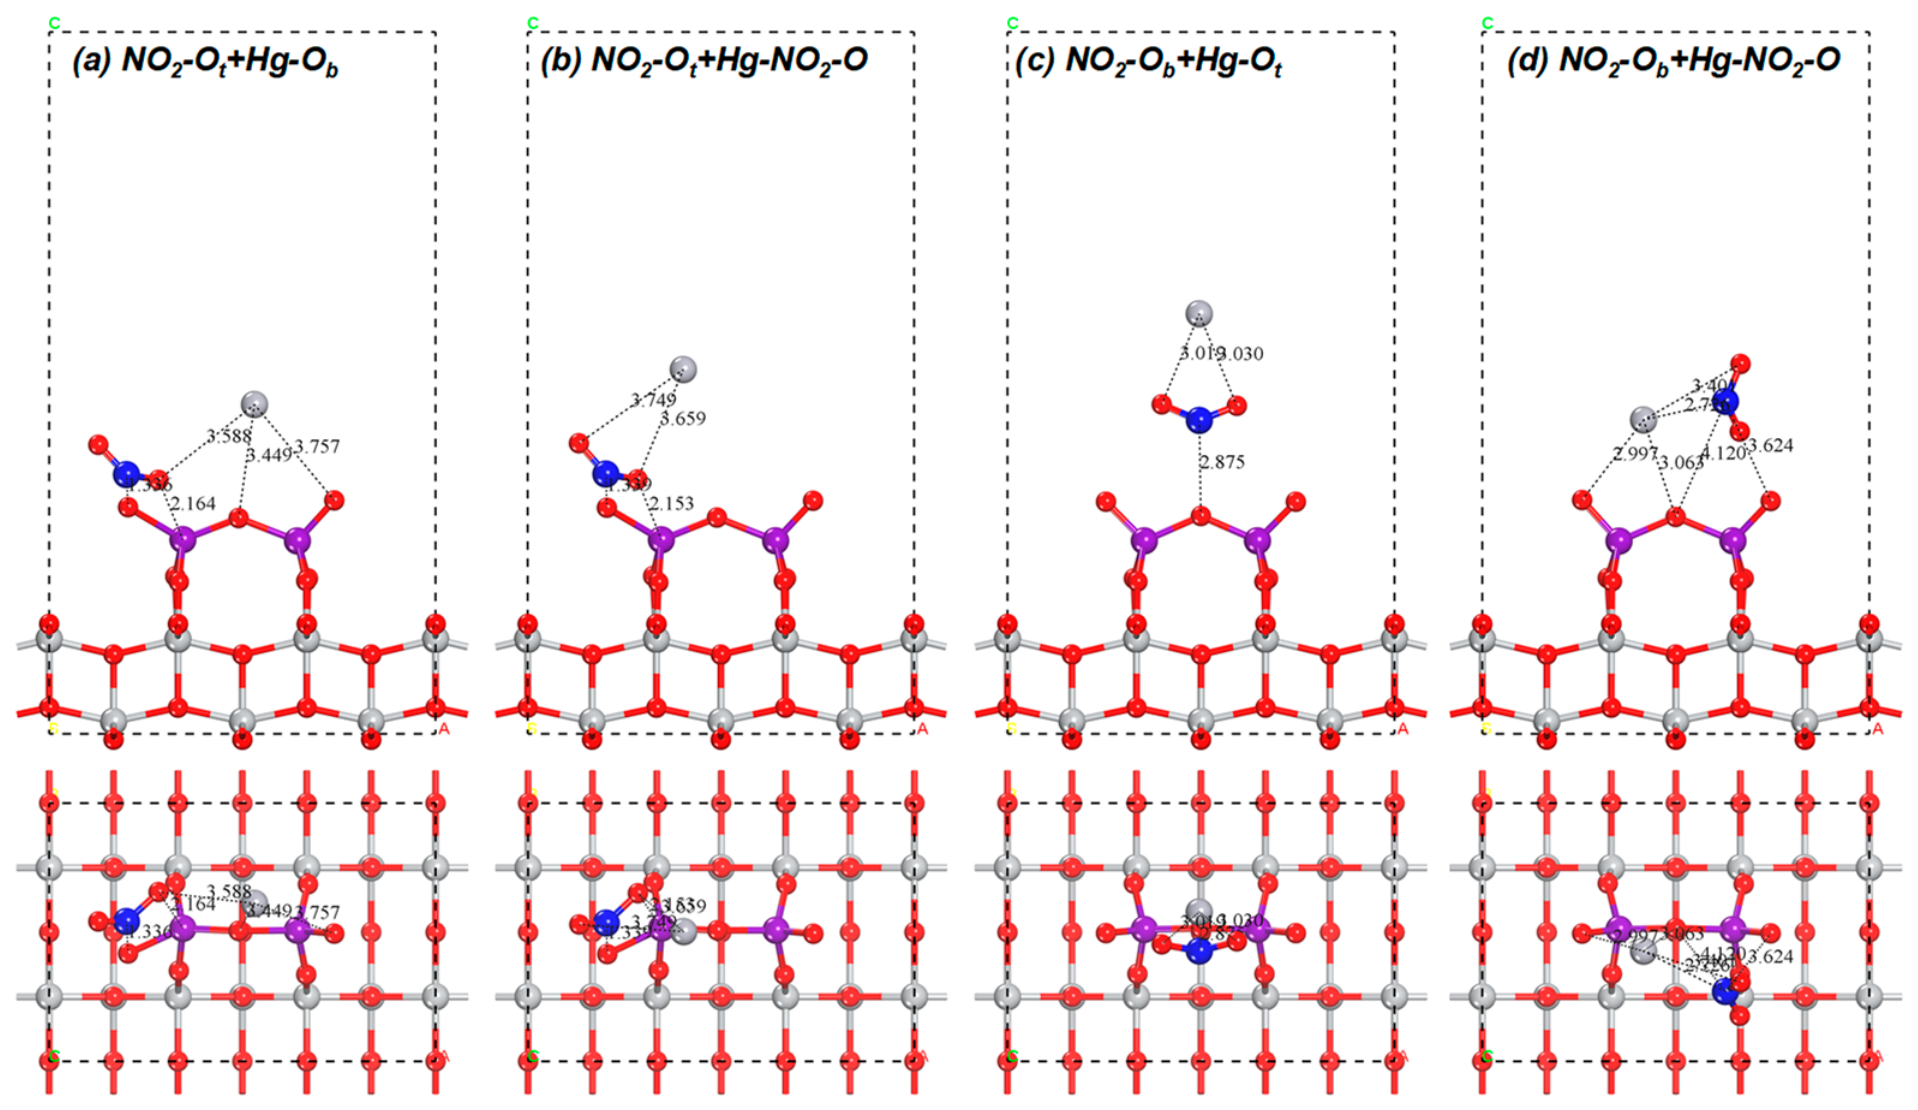
Four representative co-adsorption configurations were considered to evaluate the effect of pre-adsorbed NO2 on Hg0 adsorption over V2O5/TiO2(001), as shown in Figure 14. In the notation used here, NO2–Ot + Hg–Ob denotes a structure in which the NO2-precovered surface was first generated from the NO2–Ot adsorption model, followed by Hg0 adsorption near the Ob region. The same convention is used for the other configurations. During structural relaxation, the model initially constructed from NO2–Ot evolved into a configuration close to that obtained from the NO2–V model. Therefore, the NO2–Ot-derived precovered surface is used here to represent NO2 occupation of the V-related adsorption region, whereas the NO2–Ob model represents NO2 pre-adsorption near an oxygen-related region. Hg0 was initially placed either near a surface oxygen site, including Ot or Ob, or above the O atom of pre-adsorbed NO2, in order to compare whether Hg0 preferentially interacts with the catalyst surface or with the adsorbed NO2 moiety. The corresponding adsorption energies, characteristic distances, and Mulliken charges are listed in Table 6.
Figure 14.
Optimized configurations of Hg0 adsorbed on NO2-precovered V2O5/TiO2(001).
Table 6.
Adsorption energies, characteristic distances, and Mulliken charges for Hg0 adsorption on NO2-precovered V2O5/TiO2(001).
Compared with the clean surface, pre-adsorbed NO2 generally strengthens Hg0 adsorption. On pristine V2O5/TiO2(001), the adsorption energies of Hg–Ot and Hg–Ob are −12.31 and −12.58 kcal mol−1, respectively. After NO2 pre-adsorption, the adsorption energies decrease to −19.54 kcal mol−1 for NO2–Ot + Hg–Ob, −18.09 kcal mol−1 for NO2–Ot + Hg–NO2_O, −14.52 kcal mol−1 for NO2–Ob + Hg–Ot, and −12.27 kcal mol−1 for NO2–Ob + Hg–NO2_O. Among these configurations, NO2–Ot + Hg–Ob is the most stable, followed by NO2–Ot + Hg–NO2_O. Both are substantially more favorable than Hg0 adsorption on the clean surface. By contrast, the two NO2–Ob-based configurations show weaker stabilization, especially NO2–Ob + Hg–NO2_O, whose adsorption energy is close to those of the clean-surface cases. The optimized structures support this energy trend. For NO2–Ot + Hg–Ob, three characteristic distances are obtained: 3.588 Å, 3.449 Å, and 3.757 Å, indicating that Hg0 is stabilized around the NO2 pre-adsorbed vanadium–oxygen unit through multiple weak interactions. Similar multi-contact modes appear in NO2–Ot + Hg–NO2_O, with distances of 3.749 and 3.659 Å, respectively. Mercury is initially located above the O atom of NO2. Although the adsorption strength is slightly weaker than that of NO2–Ot + Hg–Ob, the configuration remains stabilized by interactions with the NO2-derived local environment. Nevertheless, the adsorption energy is much more negative than on a clean surface. This indicates that the O atom in the adsorbed NO2 can be used as an additional interaction site for Hg0. This behavior differs from that of NH3: NH3 mainly blocks surface adsorption sites, whereas NO2 can introduce additional oxygen-containing interaction sites and contribute to the stabilization of adsorbed Hg0. The surface based on NO2–Ob exhibits different behavior depending on the position of Hg0. In NO2–Ob + Hg–Ot, the adsorption energy is −14.52 kcal mol−1, with distances of 2.997, 3.063, and 3.401 Å. In NO2–Ob + Hg–NO2_O, the adsorption energy is −12.27 kcal mol−1, with distances of 3.019 and 3.030 Å. This is the weakest case in the NO2-precovered model. The optimized structure in Figure 14d shows that NO2 does not remain unchanged. Before Hg0 adsorption, NO2 is located above the Ob region. After the introduction of Hg0, NO2 moves to the Ot′ region. This indicates that the adsorbed NO2 species can relocate during structural relaxation when a more favorable co-adsorption configuration is formed. The Mulliken charge results show that the adsorption of Hg0 is strongly dependent on the configuration. On the clean surface, the charge of Hg-Ot and Hg-Ob to Hg0 is 0.113 e and 0.115 e, respectively. After NO2 pre-adsorption, the corresponding Hg0 charges are 0.065 e for NO2–Ot + Hg–Ob, 0.046 e for NO2–Ot + Hg–NO2_O, 0.240 e for NO2–Ob + Hg–Ot, and 0.132 e for NO2–Ob + Hg–NO2_O. Although the adsorption energy of the NO2–Ot-based configuration is more negative, its Hg0 charge transfer value is relatively small, indicating that the enhancement of its stability mainly comes from the favorable multi-center local environment, rather than large net electron transfer. In contrast, NO2–Ob + Hg-Ot combines stronger adsorption and a larger Hg0 charge, which indicates that the electron perturbation induced by Hg0 adsorption is more obvious.
Overall, pre-adsorbed NO2 strengthens Hg0 adsorption in several of the configurations examined, particularly when the V-related region is involved in the co-adsorption structure. Unlike NH3, which weakens Hg0 adsorption through site occupation and local electronic regulation, NO2 not only modifies the original V-O environment but also provides an additional O-containing interaction site for Hg0.
Hg0 Oxidative Desorption Behavior
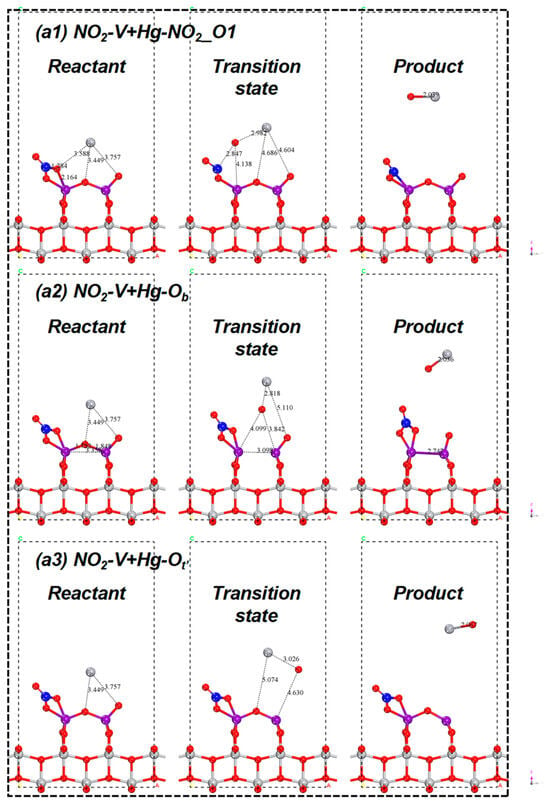
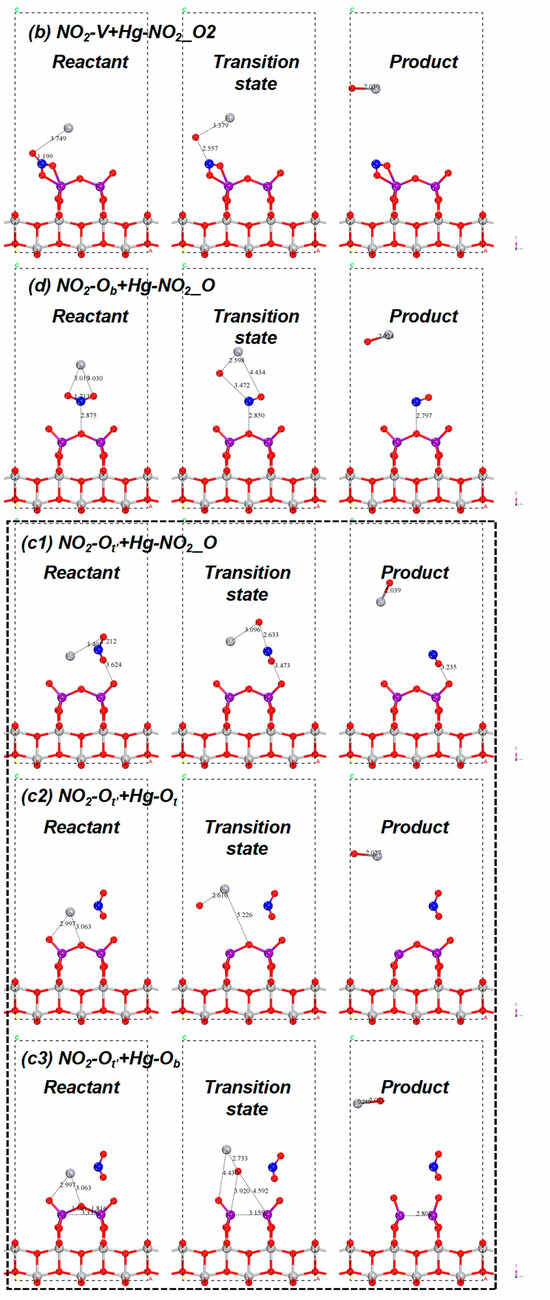
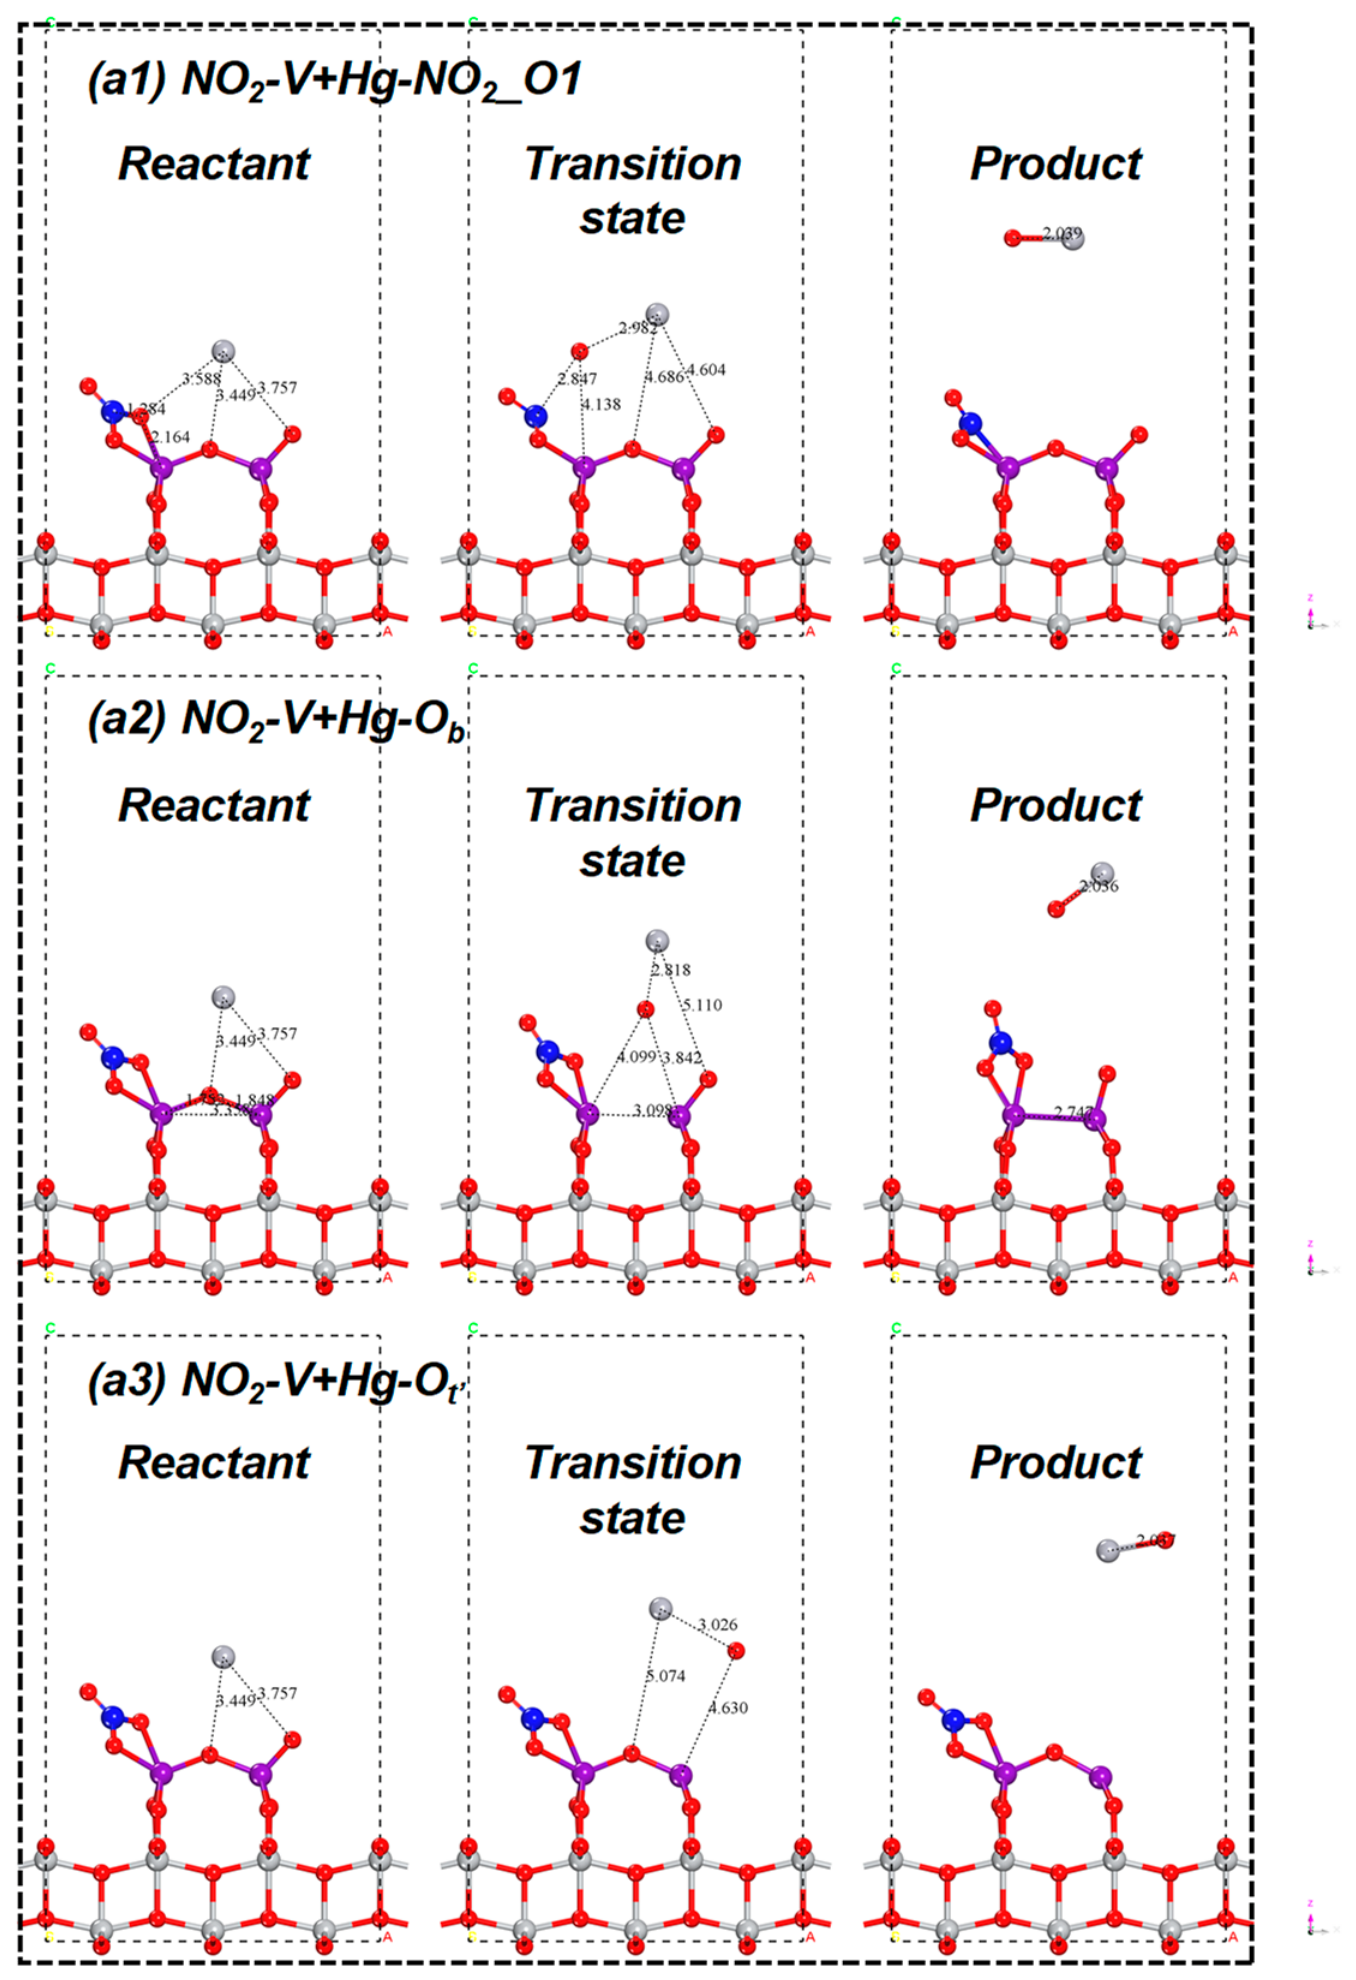
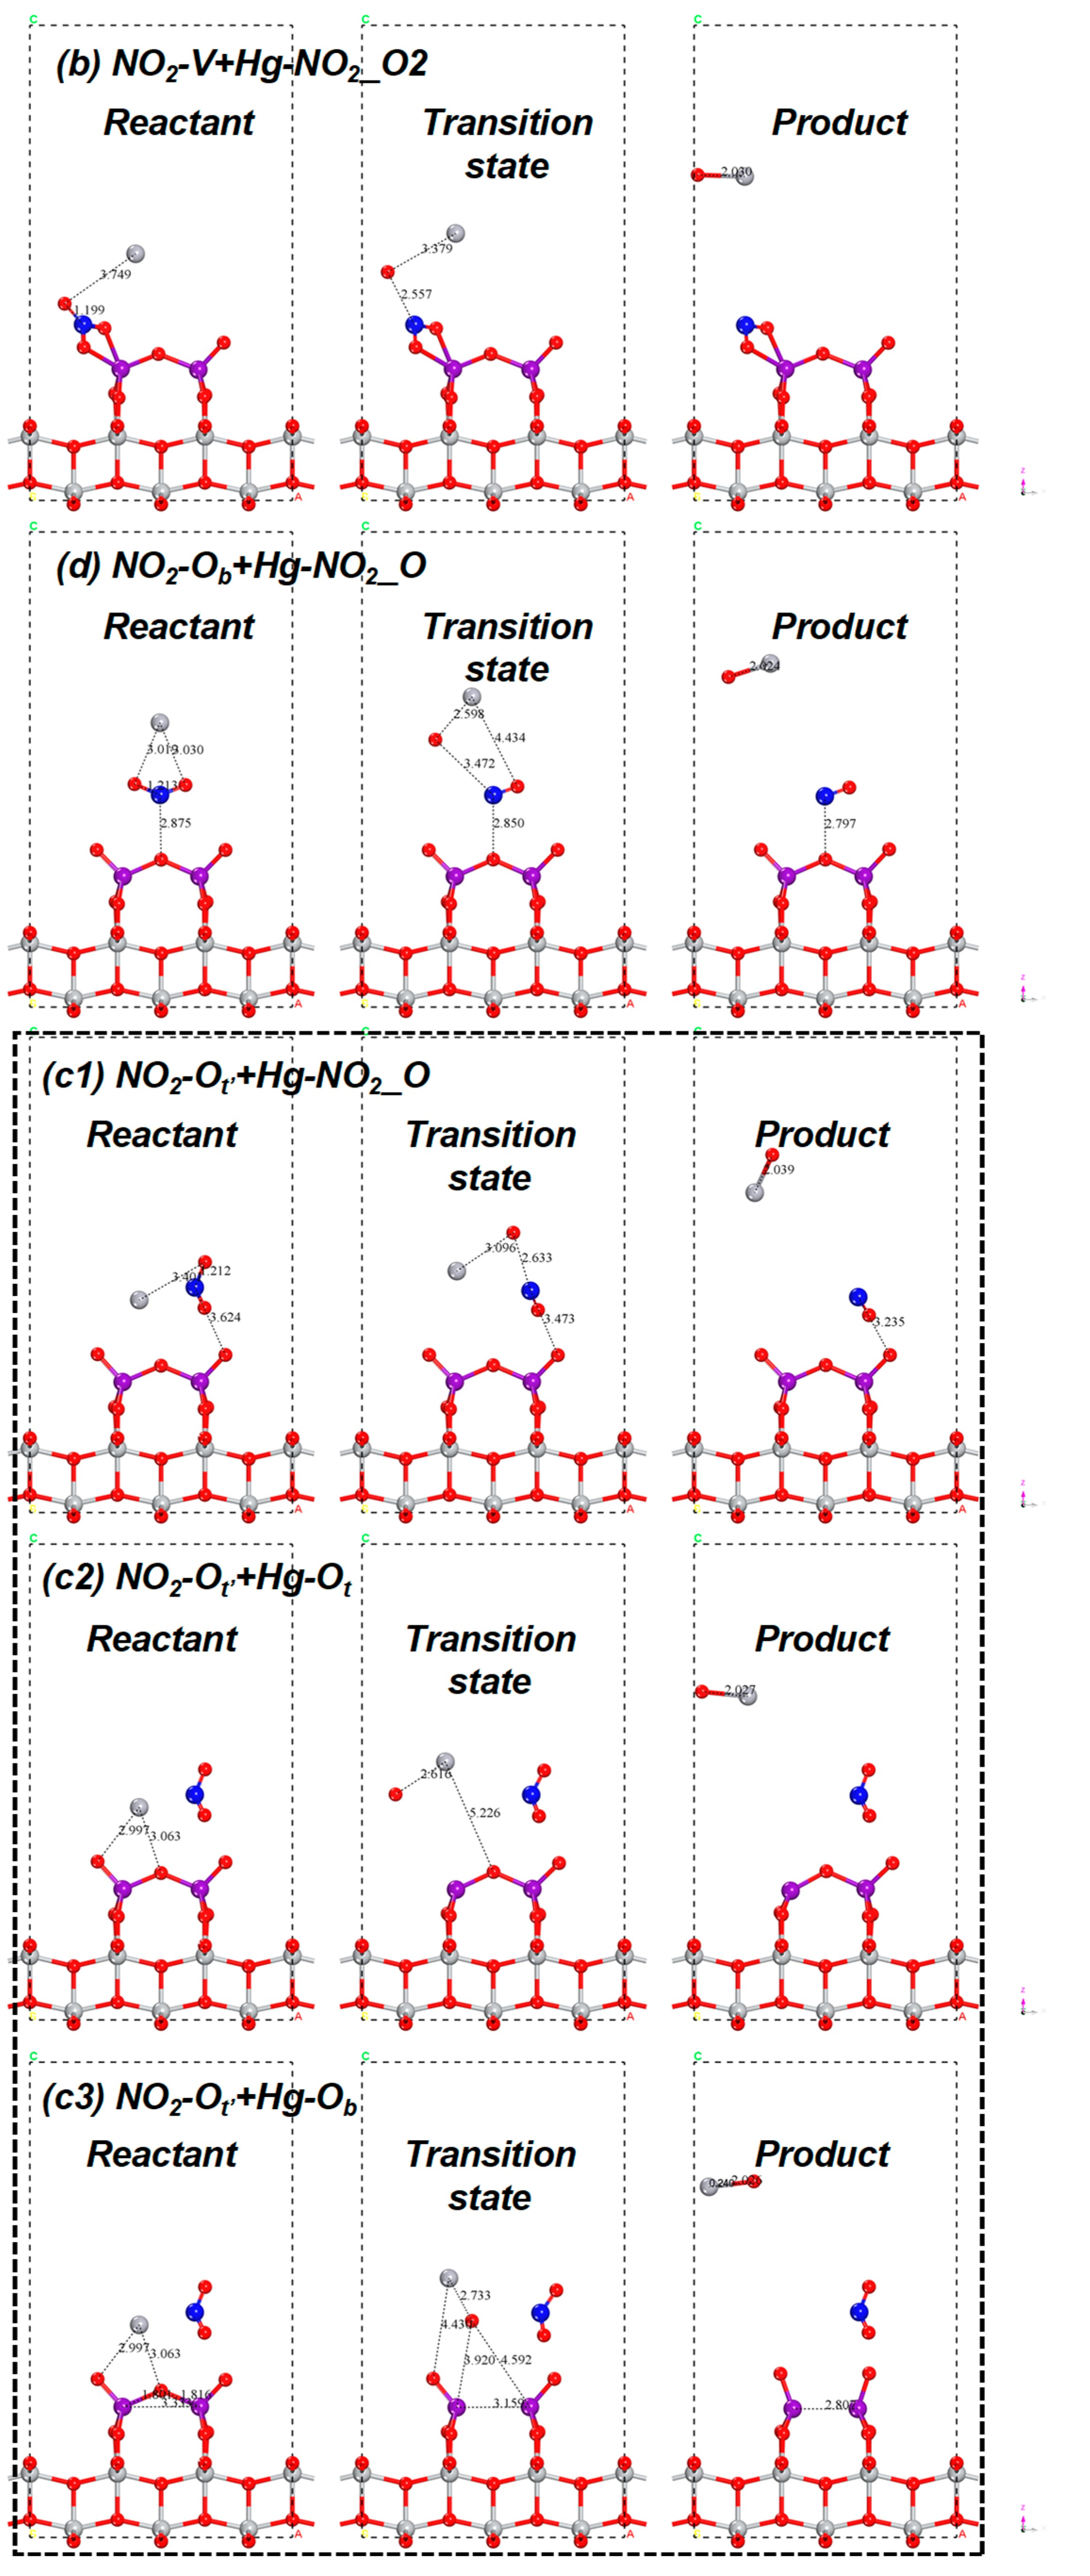
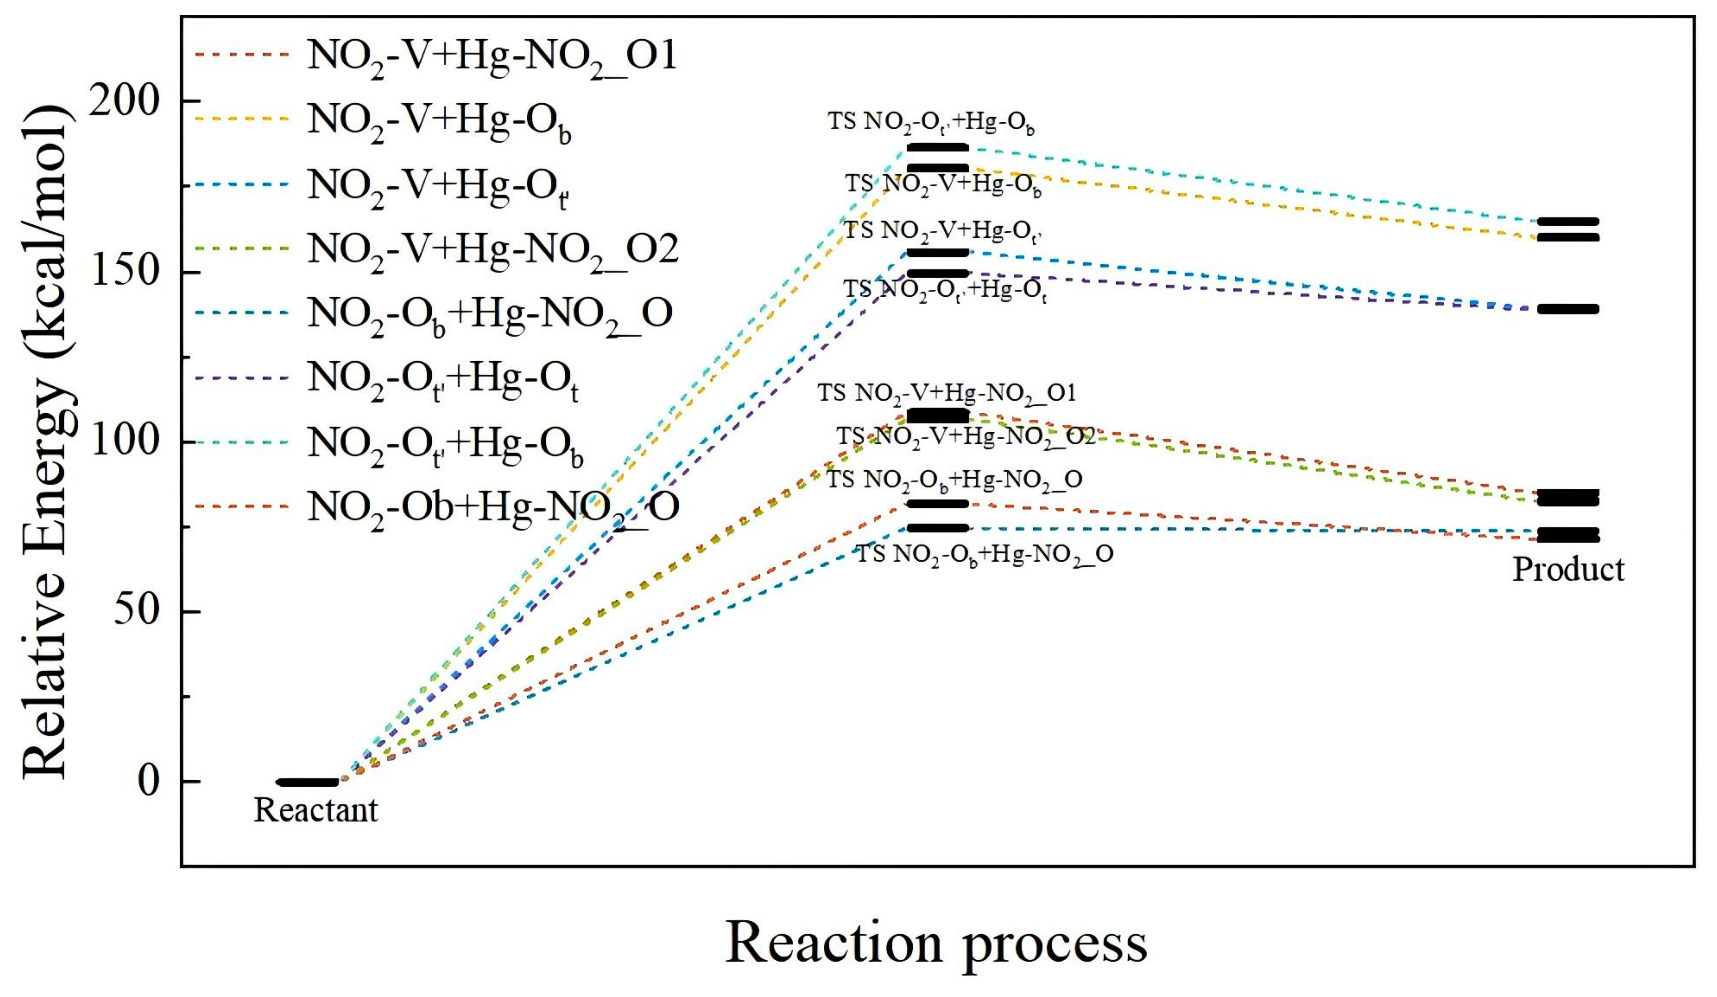
The oxidation and oxidative desorption of Hg0 on NO2-precovered V2O5/TiO2(001) surfaces were examined by transition-state calculations. The optimized structures along the reaction pathways and the corresponding energy profiles are shown in Figure 15 and Figure 16, respectively. As discussed above, pre-adsorbed NO2 can occupy the V-related region or remain near an oxygen-related region while still providing an O-containing moiety available for interaction with Hg0. Accordingly, two types of oxidation routes were compared: direct O abstraction from adsorbed NO2 and oxidation by lattice oxygen from the catalyst surface. The results show that NO2 pre-adsorption does not simply perturb the original clean-surface pathways. Instead, it introduces NO2-assisted oxygen-transfer routes with substantially lower calculated activation barriers than the lattice-oxygen-mediated pathways.
Figure 15.
Optimized reactant, transition-state, and product structures for Hg0 oxidation on NO2-precovered V2O5/TiO2(001).
Figure 16.
Reaction energy profiles for Hg0 oxidation on NO2-precovered V2O5/TiO2(001).
For the NO2–V-precovered surface, four representative pathways were considered. In the NO2–V + Hg–NO2_O1 and NO2–V + Hg–NO2_O2 pathways, Hg0 reacts directly with an O atom of the adsorbed NO2 species to form an HgO-like product. The corresponding reaction and activation energies are 84.943 and 108.545 kcal mol−1, respectively. By contrast, when Hg0 reacts with lattice oxygen on the same NO2–V precovered surface, the energetic cost increases markedly. For the NO2–V + Hg–Ot′ pathway, the reaction and activation energies are 139.116 and 155.786 kcal mol−1, respectively, while for the NO2–V + Hg–Ob pathway, they rise further to 160.037 and 180.331 kcal mol−1. These results show that oxygen transfer from adsorbed NO2 has a lower calculated activation barrier than lattice-oxygen extraction within the same computational framework, with a barrier reduction of approximately 47–74 kcal mol−1. A similar pattern appears on the NO2–Ot′-precovered surface. Three pathways were examined, which involve oxygen transfer from NO2, from a nearby Ot atom, and from an Ob atom. The NO2–Ot′ + Hg–NO2_O pathway has the lowest calculated activation barrier among these three pathways, with a reaction energy of 73.672 kcal mol−1 and an activation barrier of 74.581 kcal mol−1. The lattice-oxygen pathways require much higher energies. NO2–Ot′ + Hg–Ot has reaction and activation energies of 138.676 and 149.585 kcal mol−1, and NO2–Ot′ + Hg–Ob requires 164.750 and 186.384 kcal mol−1. The difference in activation energy further indicates that oxygen transfer from adsorbed NO2 is energetically preferred over direct extraction of lattice oxygen among the pathways considered. For the NO2–Ob + Hg–NO2_O pathway, the O atom of the pre-adsorbed NO2 species directly serves as the oxygen donor for Hg0 oxidation. The reaction energy is 71.346 kcal mol−1, and the activation energy is 81.767 kcal mol−1, both of which are considerably lower than those associated with lattice-oxygen-mediated oxidation on either the clean or NO2-precovered surfaces. During the oxygen-transfer step, Hg0 approaches one O atom of the adsorbed NO2 species. In the optimized product state, this O atom is transferred to Hg, forming a detached HgO-like moiety, whereas the N atom and the remaining O atom of the original NO2 remain associated with the catalyst surface as an NO-like residue (denoted as NO *). Accordingly, the elementary step can be represented schematically as Hg0 + NO2 * → HgO (detached) + NO *. Here, NO * denotes the immediate surface-associated residue identified in the optimized product structure; its independent adsorption stability and subsequent desorption or further reaction were not evaluated in the present study. The present pathway therefore describes the NO2-to-Hg oxygen-transfer step rather than the complete subsequent NOx reaction cycle. In this work, “oxidative desorption” refers to the coupled oxygen-transfer and product-detachment process observed along the calculated pathway; a separate HgO desorption barrier was not calculated. This structural evolution shows that the O atom of adsorbed NO2 can directly participate in the oxygen-transfer step, regardless of whether NO2 is initially located near a V-related site or an oxygen-related site.
A comparison with the clean surface further highlights the relative energetic effect of NO2 pre-adsorption. On clean V2O5/TiO2(001), Hg0 oxidation proceeds only through lattice-oxygen abstraction and requires high barriers of 146.817, 177.851, and 184.529 kcal mol−1 for the Ot, Ob, and interfacial pathways, respectively. After NO2 pre-adsorption, the NO2-assisted pathways reduce the barriers to 74.581–108.545 kcal mol−1, whereas lattice-oxygen oxidation on the same NO2-precovered surfaces retains substantially higher calculated barriers (Table 7). Therefore, NO2 pre-adsorption does not simply modify the energetics of the original lattice-oxygen pathway; it also introduces an alternative oxygen-transfer channel in which the adsorbed NO2 species directly participates. This alternative oxygen-transfer pathway provides a possible mechanistic interpretation of the NO2-associated promotion of Hg0 oxidation reported experimentally under fast-SCR conditions [7].
Table 7.
Reaction energies and activation barriers for Hg0 oxidation on NO2-precovered V2O5/TiO2(001).
3.4. Effect of Co-Pre-Adsorbed NH3 and NO2 on Hg0 Oxidation over V2O5/TiO2(001)
3.4.1. Hg0 Adsorption Behavior
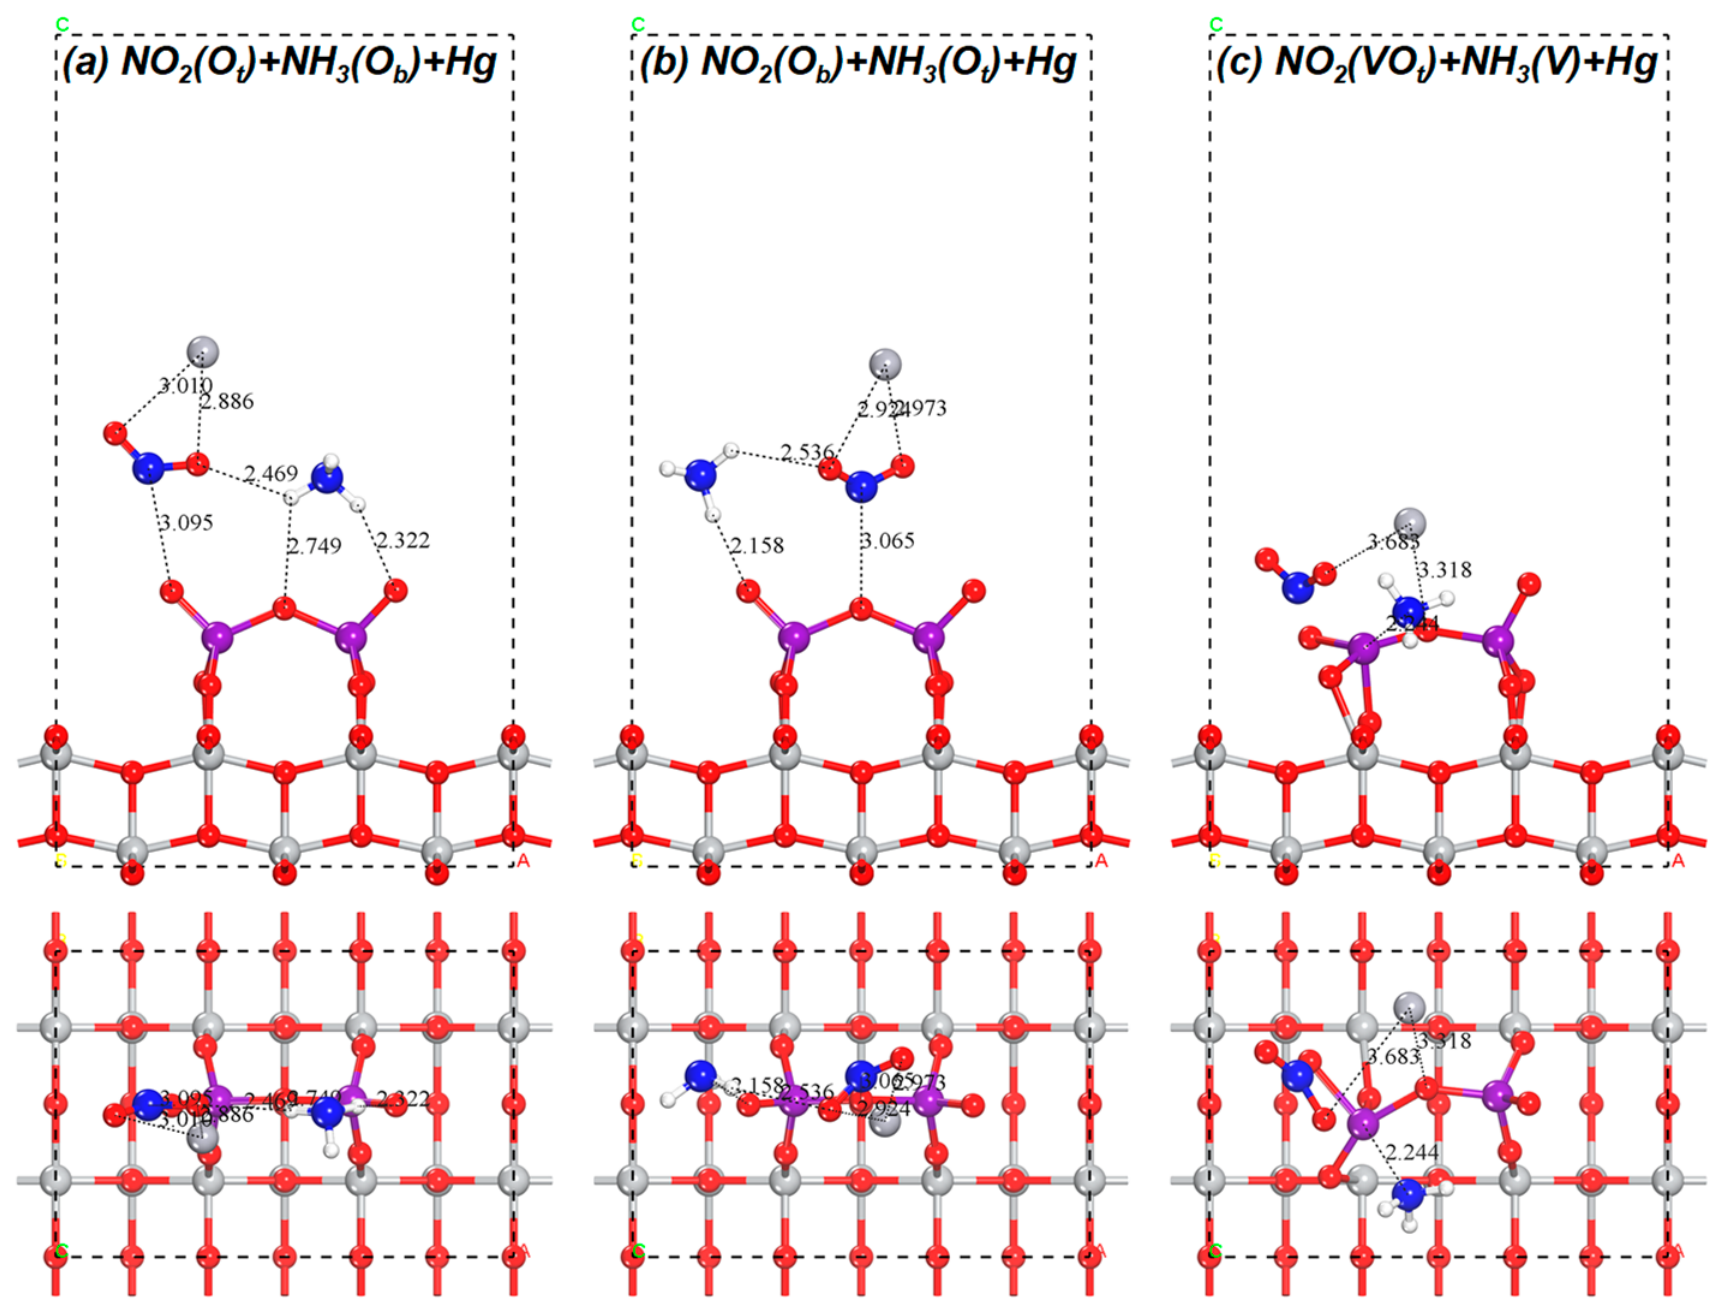
Hg0 adsorption was examined on NH3/NO2 co-precovered V2O5/TiO2(001) surfaces to clarify how the coexistence of the two adsorbates affects the initial capture of Hg0. As shown in Figure 17, two types of co-precovered surfaces were considered. In the first type, NH3 and NO2 occupy oxygen-related adsorption regions that could otherwise serve as possible Hg0 adsorption sites, represented by NO2(Ot) + NH3(Ob) + Hg and NO2(Ob) + NH3(Ot) + Hg. In the second type, both adsorbates are associated with the V-related region, represented by NO2(VOt) + NH3(V) + Hg. This configuration was used to examine the effect of simultaneous modification of the V-centered local environment on Hg0 adsorption. In all models, Hg0 was initially placed above the NO2-related region or a neighboring surface region before full structural optimization. The corresponding adsorption energies, representative interatomic distances, and Mulliken charges are listed in Table 8.
Figure 17.
Optimized configurations of Hg0 adsorbed on NH3/NO2-co-precovered V2O5/TiO2(001).
Table 8.
Adsorption energies, characteristic distances, and Mulliken charges for Hg0 adsorption on NH3/NO2-co-precovered V2O5/TiO2(001).
The adsorption energy results show that the coexistence of NO2 and NH3 does not lead to the strong inhibition of Hg0 adsorption observed on NH3-precovered surfaces. On clean V2O5/TiO2(001), Hg–Ot and Hg–Ob have adsorption energies of −12.31 and −12.58 kcal mol−1, respectively. After NH3 pre-adsorption, Hg0 adsorption becomes weaker, with adsorption energies of −11.04 to −11.84 kcal mol−1. In contrast, on the NH3/NO2 co-precovered surface, the adsorption energies are −12.606, −12.723, and −15.469 kcal mol−1, corresponding to NO2(Ot) + NH3(Ob) + Hg0, NO2(Ob) + NH3(Ot) + Hg0, and NO2(VOt) + NH3(V) + Hg0, respectively. The first two adsorption energies are comparable to those on the clean surface, or even slightly more negative, whereas NO2(VOt) + NH3(V) + Hg0 shows significantly stronger stability. This indicates that NO2 can partially counteract the inhibitory effect of NH3 on Hg0 adsorption. The optimized geometries provide further evidence for this role of NO2. For NO2(Ot) + NH3(Ob) + Hg0, the two Hg–O distances of 3.010 and 2.886 Å correspond to interactions between Hg0 and the two O atoms of pre-adsorbed NO2. A similar feature is found for NO2(Ob) + NH3(Ot) + Hg0, where the optimized Hg–O distances of 2.924 and 2.937 Å also involve the O atoms of the adsorbed NO2 species. In both configurations, Hg0 remains close to the NO2-derived O-containing region rather than being displaced away from the surface. Thus, once NO2 is present, its O atoms can act as accessible local interaction sites for Hg0 adsorption. For NO2(VOt) + NH3(V) + Hg0, the optimized distances of 3.683 and 3.318 Å correspond to the separations between Hg0 and the O atom of pre-adsorbed NO2, and between Hg0 and a neighboring surface O atom in the V-centered region, respectively. Unlike the oxygen-region co-precovered models, Hg0 in this configuration is not stabilized by a single isolated site. In contrast, adsorption is driven by a combined local environment that involves the adsorbed NO2 species and the nearby surface oxygen. Therefore, the strong adsorption energy of −15.469 kcal mol−1 can be attributed to the synergistic multi-center adsorption environment formed around the V-related region.
The Mulliken charge analysis further highlights the configuration dependence of Hg0 stability. The calculated Hg0 charges are 0.145 e, 0.136 e, and 0.037 e, corresponding to NO2(Ot) + NH3(Ob) + Hg0, NO2(Ob) + NH3(Ot) + Hg0, and NO2(VOt) + NH3(V) + Hg0, respectively. The first two values are higher than those obtained on the NH3 pre-adsorbed surface and slightly higher than those obtained on the clean surface, indicating that when NH3 and NO2 occupy the adjacent oxygen-related regions, Hg0 will still undergo significant electronic perturbation due to adsorption. In contrast, NO2(VOt) + NH3(V) + Hg0 has the smallest Hg0 charge but the most favorable adsorption energy.
In general, the role of NO2 in the NH3/NO2 co-precovered system is significantly different from that of NH3 alone. In the case of NH3 alone, the pre-adsorbed NH3 mainly occupies the catalyst-related adsorption region, which reduces the accessibility of Hg adsorption sites. However, in the co-precovered system, the O atom of adsorbed NO2 provides additional interaction sites for Hg0. Therefore, although NH3 still occupies part of the surface adsorption environment, the presence of NO2 introduces an oxygen-containing local environment that can still be in contact with Hg, which helps to maintain or even enhance the initial adsorption of Hg0 on V2O5/TiO2(001).
3.4.2. Hg0 Oxidative Desorption Behavior
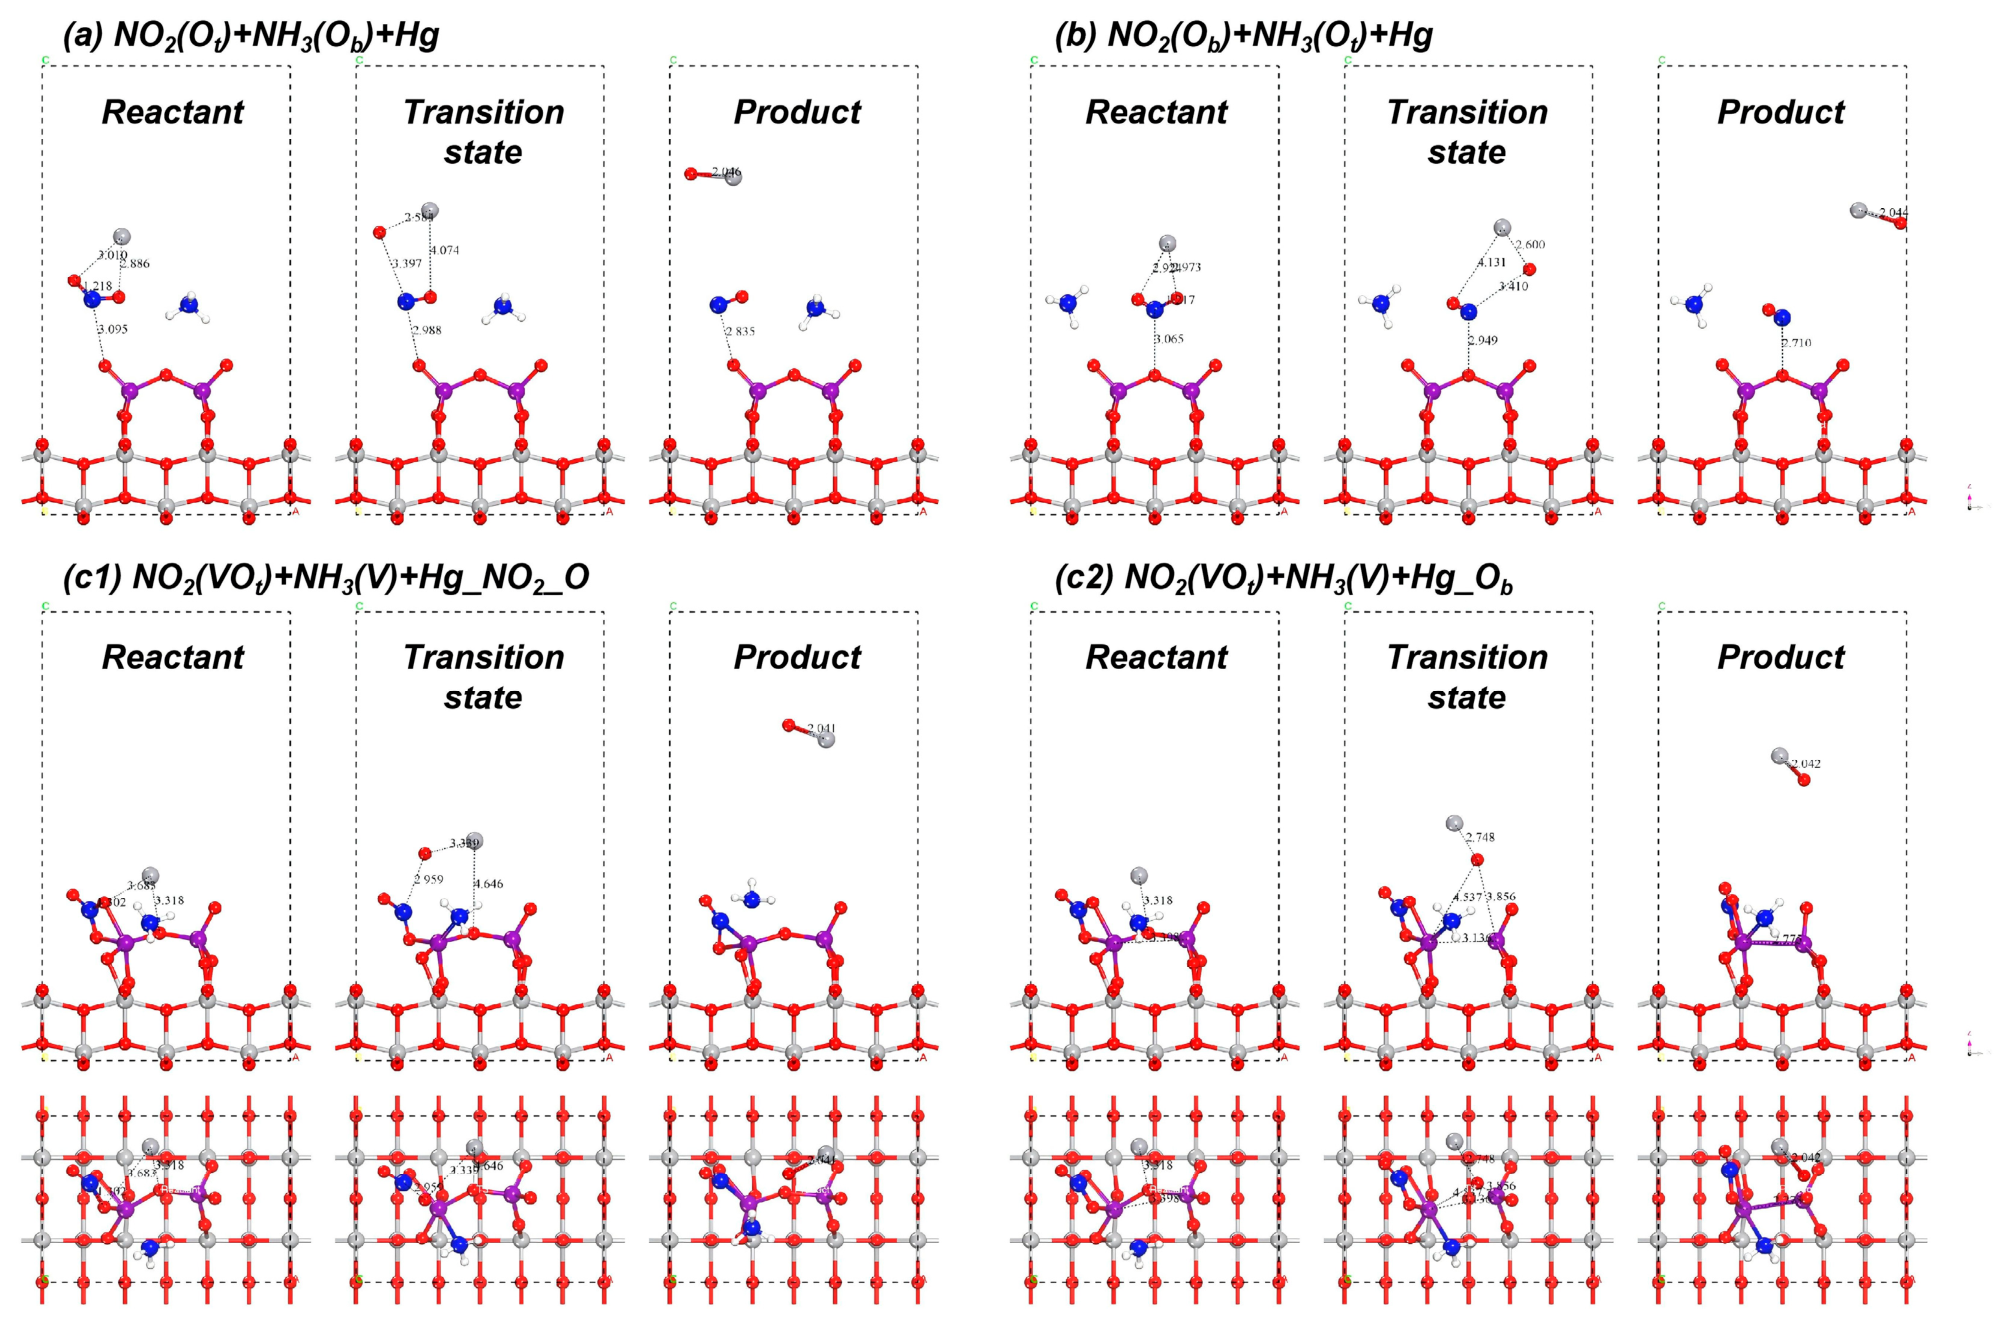
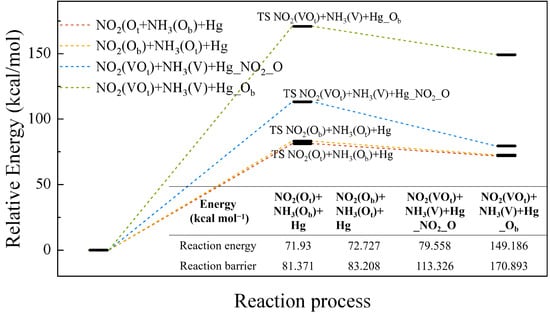
Three representative oxidation pathways were calculated on the NH3/NO2 co-covered V2O5/TiO2(001) surface to evaluate how the coexistence of NH3/NO2 affects the subsequent Hg0 conversion. The optimized structures along the reaction coordinates are shown in Figure 18, and the corresponding reaction energy profiles are presented in Figure 19. In the first two configurations, NH3 and NO2 occupy neighboring oxygen-related regions, represented by NO2(Ot) + NH3(Ob) + Hg0 and NO2(Ob) + NH3(Ot) + Hg0. In the third configuration, both adsorbates are associated with the V-centered local environment, represented by NO2(VOt) + NH3(V) + Hg0. For these surfaces, two types of oxidation routes were examined: oxygen transfer from the adsorbed NO2 species and oxidation by lattice oxygen from the catalyst surface. The calculated results show that, even under NH3/NO2 co-precovered conditions, oxidation through pre-adsorbed NO2 exhibits a lower calculated activation barrier than lattice-oxygen abstraction within the present computational framework.
Figure 18.
Optimized reactant, transition-state, and product structures for Hg0 oxidation on NH3/NO2-co-precovered V2O5/TiO2(001).
Figure 19.
Reaction energy profiles for Hg0 oxidation on NH3/NO2-co-precovered V2O5/TiO2(001).
For NO2(Ot) + NH3(Ob) + Hg and NO2(Ob) + NH3(Ot) + Hg, Hg0 is oxidized by an O atom of the adsorbed NO2 species, as shown in Figure 18a,b. The corresponding reaction energies and activation barriers are 71.930/81.371 kcal mol−1 and 72.727/83.208 kcal mol−1, respectively. The two pathways are close in energy, showing that when NH3 and NO2 occupy neighboring oxygen-related regions, the initial location of NO2 near Ot or Ob has only a limited influence on the overall oxidation energetics. In both cases, Hg0 approaches the O atom of pre-adsorbed NO2, followed by oxygen transfer and formation of a detached HgO-like product, while the N atom and the remaining O atom of the original NO2 remain associated with the surface as an NO-like residue (NO *). The subsequent fate of this NO * species was not considered in the present elementary-step analysis. These barriers remain close to those of the most favorable NO2-only precovered routes, which means that NH3 does not significantly suppress the NO2-assisted oxidation channel when it is located away from the V-centered adsorption environment. A different behavior is observed for the NO2(VOt) + NH3(V) + Hg co-precovered surface, where both adsorbates modify the V-centered local environment. Two oxidation pathways were examined, as shown in Figure 18(c1,c2). In NO2(VOt) + NH3(V) + Hg_NO2_O, Hg0 is oxidized by the O atom of pre-adsorbed NO2, giving a reaction energy of 79.558 kcal mol−1 and an activation barrier of 113.326 kcal mol−1. In contrast, when Hg0 reacts with the lattice Ob atom in NO2(VOt) + NH3(V) + Hg_Ob, the reaction energy and activation barrier increase sharply to 149.186 and 170.893 kcal mol−1, respectively. Thus, even when NH3 and NO2 are both concentrated around the V-centered region, oxygen transfer from adsorbed NO2 remains energetically preferred over the corresponding lattice-oxygen pathway. The barrier difference between the NO2-assisted and lattice-oxygen routes exceeds 57 kcal mol−1, indicating a relative energetic preference for oxygen transfer from pre-adsorbed NO2 over the lattice-oxygen pathway.
Compared to the clean surface and NH3 precovered surface, the NO2-assisted pathway under NH3/NO2 co-coverage maintains a lower energy barrier. In particular, the two oxygen-region co-precovered pathways in Figure 18a,b exhibit calculated barriers of approximately 81–83 kcal mol−1, which are lower than the corresponding lattice-oxygen-mediated barriers within the present computational framework. However, when NH3 and NO2 simultaneously occupy the V-centered local environment, the barrier of the NO2-assisted pathway rises to 113.326 kcal mol−1. This value is higher than those of the most favorable NO2-only pathways, indicating that NH3 increases the energetic cost of the NO2-assisted pathway when both adsorbates modify the same local V–O environment.
Overall, the co-precovered systems show a competitive but asymmetric effect between NH3 and NO2. NH3 increases the calculated barrier of the NO2-assisted pathway when both adsorbates perturb the V-centered region, but NO2-derived oxygen transfer remains energetically preferred over the corresponding lattice-oxygen pathway. Among the pathways examined, the lower-energy oxidation route still involves the O atom of pre-adsorbed NO2 rather than lattice oxygen. Therefore, within the present computational model, NO2-derived oxygen provides the relatively lower-energy oxidation route, whereas NH3 mainly modifies the local adsorption environment and can increase the calculated barrier when both species perturb the same V-centered region. These configuration-dependent effects are qualitatively consistent with previous experimental observations of NH3 inhibition and NO2-associated promotion of Hg0 oxidation [7,12].
4. Conclusions
In this work, DFT calculations were performed using a V2O5/TiO2(001) model to clarify the effects of representative NH3- and NO2-derived surface environments on Hg0 adsorption and oxidation relevant to FSCR conditions. On the clean surface, Hg0 adsorption was generally weak. Although the Ob′ site showed the strongest adsorption energy of −21.22 kcal mol−1, it did not provide the most favorable oxidation route. Oxidation by lattice oxygen remained energetically demanding, with calculated barriers of 146.817, 177.851, and 184.529 kcal mol−1 for the Ot, Ob, and Ob′ pathways, respectively. NH3 preferentially adsorbs at exposed V-related sites through N coordination, particularly in the V–Ot local environment. Its pre-adsorption weakens Hg0 capture, as reflected by the less negative adsorption energies, whereas the subsequent lattice-oxygen-mediated oxidation pathways remain energetically similar to those on the clean surface. Therefore, NH3 mainly affects Hg0 conversion at the initial adsorption stage by modifying or occupying the local adsorption environment. NO2 pre-adsorption produces a different energetic response. In addition to modifying Hg0 adsorption, pre-adsorbed NO2 introduces alternative oxygen-transfer pathways with calculated activation barriers of 74.581–108.545 kcal mol−1, substantially lower than those of direct lattice-oxygen abstraction within the same computational framework. Under NH3/NO2 co-precoverage, oxygen transfer from NO2 remains the relatively lower-energy route among the pathways examined. When NH3 and NO2 occupy neighboring oxygen-related regions, the calculated barriers are approximately 81–83 kcal mol−1, whereas simultaneous perturbation of the V-centered local environment increases the NO2-assisted barrier to 113.326 kcal mol−1. These results indicate that NH3 can modify the relative energetics of the NO2-assisted pathway. Overall, the present calculations indicate that NH3 and NO2 exert different effects on Hg0 conversion on V2O5/TiO2(001): NH3 mainly weakens the formation of favorable Hg0 adsorption precursors, whereas pre-adsorbed NO2 introduces an alternative oxygen-transfer channel that is relatively lower in energy than direct lattice-oxygen oxidation among the pathways examined in the present model. It should be noted that, since these calculations are based on a simplified surface model without thermal and entropic corrections, the results should be interpreted as relative energetic trends rather than quantitative predictions of reaction kinetics under practical FSCR conditions.
Author Contributions
Conceptualization, X.D.; methodology, X.D.; software, X.D.; validation, Q.R.; formal analysis, G.X.; investigation, H.W.; resources, X.D.; data curation, J.L. and N.G.; writing—original draft preparation, H.W.; writing—review and editing, J.L., G.X., S.D., Z.C., N.G., and X.D.; visualization, S.D.; supervision, X.D.; project administration, Z.C.; funding acquisition, X.D. All authors have read and agreed to the published version of the manuscript.
Funding
This work was supported by the Project on Mercury Emission Characteristics and Operational Optimization Testing of Coal-Fired Boilers (GSKJ-25-50).
Data Availability Statement
The data presented in this study are available on request from the corresponding author. The data are not publicly available due to the data forming part of an ongoing study.
Conflicts of Interest
Hu Wang, Jiao Liu, Guowei Xie and Shoubao Duan were employed by CHN Energy Guoyuan Electric Power (Beijing) Co., Ltd. Zhiyuan Cheng and Ninglu Geng were employed by the National Environmental Protection Research Institute for Electric Power Co., Ltd. The remaining authors declare that the research was conducted in the absence of any commercial or financial relationships that could be construed as a potential conflict of interest. The funder was not involved in the study design; the collection, analysis, or interpretation of data; the writing of this article; or the decision to submit it for publication.
References
- Chen, C.; Cao, Y.; Liu, S.; Chen, J.; Jia, W. Review on the latest developments in modified vanadium-titanium-based SCR catalysts. Chin. J. Catal. 2018, 39, 1347–1365. [Google Scholar] [CrossRef] [Scilit]
- Wang, H.; Yuan, B.; Hao, R.; Zhao, Y.; Wang, X. A critical review on the method of simultaneous removal of multi-air-pollutant in flue gas. Chem. Eng. J. 2019, 378, 122155. [Google Scholar] [CrossRef] [Scilit]
- Balasundaram, K.; Sharma, M. Technology for mercury removal from flue gas of coal based thermal power plants: A comprehensive review. Crit. Rev. Environ. Sci. Technol. 2019, 49, 1700–1736. [Google Scholar] [CrossRef] [Scilit]
- Ishag, A.; Yue, Y.; Xiao, J.; Huang, X.; Sun, Y. Recent advances on the adsorption and oxidation of mercury from coal-fired flue gas: A review. J. Clean. Prod. 2022, 367, 133111. [Google Scholar] [CrossRef] [Scilit]
- He, C.; Shen, B.; Chi, G.; Li, F. Elemental mercury removal by CeO2/TiO2-PILCs under simulated coal-fired flue gas. Chem. Eng. J. 2016, 300, 1–8. [Google Scholar] [CrossRef] [Scilit]
- Chen, C.; Jia, W.; Liu, S.; Cao, Y. Simultaneous NO Removal and Hg0 Oxidation over CuO Doped V2O5-WO3/TiO2 Catalysts in Simulated Coal-Fired Flue Gas. Energy Fuels 2018, 32, 7025–7034. [Google Scholar] [CrossRef] [Scilit]
- Jia, W.; Wang, Y.; Ling, S.; He, L.; Chen, B.; Liu, S.; Chen, C.; Cao, Y. Synergetic NOx Removal and Hg0 Oxidation over V2O5–WO3/TiO2 Catalysts in a Simulated Fast SCR Atmosphere: Catalytic Performance and Hg0 Oxidation Mechanism. Energy Fuels 2024, 38, 8035–8043. [Google Scholar] [CrossRef] [Scilit]
- Kamata, H.; Ueno, S.-I.; Naito, T.; Yukimura, A. Mercury oxidation over the V2O5 (WO3)/TiO2 commercial SCR catalyst. Ind. Eng. Chem. Res. 2008, 47, 8136–8141. [Google Scholar] [CrossRef] [Scilit]
- Yang, J.; Yang, Q.; Sun, J.; Liu, Q.; Zhao, D.; Gao, W.; Liu, L. Effects of mercury oxidation on V2O5–WO3/TiO2 catalyst properties in NH3-SCR process. Catal. Commun. 2015, 59, 78–82. [Google Scholar] [CrossRef] [Scilit]
- Liu, R.; Xu, W.; Tong, L.; Zhu, T. Role of NO in Hg0 oxidation over a commercial selective catalytic reduction catalyst V2O5–WO3/TiO2. J. Environ. Sci. 2015, 38, 126–132. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Shi, J.; Chen, J.; Wang, J.; Yin, R.; Wang, B.; Xiong, S.; Wang, Z.; Liu, H.; Li, J. Vanadium-density-dependent reactivity for simultaneous removal of NOx and Hg0 over V2O5/TiO2 catalyst. Fuel 2023, 332, 126189. [Google Scholar] [CrossRef] [Scilit]
- Park, J.H.; Nguyen, T.P.T.; Kim, M.H.; Hong, Y. Effect of Vanadium Oxide Loading for the Simultaneous NO Reduction and Hg0 Oxidation over a V2O5-WO3/TiO2 Catalyst. Ind. Eng. Chem. Res. 2023, 62, 21599–21608. [Google Scholar] [CrossRef] [Scilit]
- Suarez Negreira, A.; Wilcox, J. DFT study of Hg oxidation across vanadia-titania SCR catalyst under flue gas conditions. J. Phys. Chem. C 2013, 117, 1761–1772. [Google Scholar] [CrossRef] [Scilit]
- Gao, Y.; Li, Z. A DFT study of the Hg0 oxidation mechanism on the V2O5-TiO2 (001) surface. Mol. Catal. 2017, 433, 372–382. [Google Scholar] [CrossRef] [Scilit]
- Gao, Y.; Li, Z.; Hao, Y. Effect of M-doped (M=Cr, Fe, Co, and Nb) V2O5/TiO2 (001) on mercury oxidation: The insights from DFT calculation. J. Phys. Chem. C 2017, 121, 27963–27975. [Google Scholar]
- Wu, Y.-w.; Ali, Z.; Lu, Q.; Liu, J.; Xu, M.-x.; Zhao, L.; Yang, Y.-p. Effect of WO3 doping on the mechanism of mercury oxidation by HCl over V2O5/TiO2 (001) surface: Periodic density functional theory study. Appl. Surf. Sci. 2019, 487, 369–378. [Google Scholar] [CrossRef] [Scilit]
- Su, J.; Yang, J.; Zhang, M.; Gao, M.; Zhang, Y.; Gao, M.; Chen, L.; Huang, Y.; Wang, Z.; Shen, B. Mechanism of Mo and Sb species improving Hg0 oxidation performance of V2O5/TiO2 catalyst: Density function theory study. Appl. Surf. Sci. 2023, 617, 156612. [Google Scholar] [CrossRef] [Scilit]
- Su, J.; Yang, J.; Zhang, M.; Gao, M.; Zhang, Y.; Gao, M.; Chen, L.; Huang, Y.; Wang, Z.; Shen, B. Improvement mechanism of Ru species on Hg0 oxidation reactivity over V2O5/TiO2 Catalyst: A density functional theory study. Chem. Eng. Sci. 2023, 274, 118689. [Google Scholar] [CrossRef] [Scilit]
- Zhang, B.; Liu, J.; Dai, G.; Chang, M.; Zheng, C. Insights into the mechanism of heterogeneous mercury oxidation by HCl over V2O5/TiO2 catalyst: Periodic density functional theory study. Proc. Combust. Inst. 2015, 35, 2855–2865. [Google Scholar] [CrossRef] [Scilit]
- Yang, H.G.; Sun, C.H.; Qiao, S.Z.; Zou, J.; Liu, G.; Smith, S.C.; Cheng, H.M.; Lu, G.Q. Anatase TiO2 single crystals with a large percentage of reactive facets. Nature 2008, 453, 638–641. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Mi, T.-G.; Wu, Y.-W.; Zhou, X.-Y.; Li, W.-T.; Zhao, L.; Liu, J.; Lu, Q. Catalytic oxidation of CO over V2O5/TiO2 and V2O5-WO3/TiO2 catalysts: A DFT study. Fuel Process. Technol. 2021, 213, 106678. [Google Scholar] [CrossRef] [Scilit]
- Calatayud, M.; Tielens, F.; De Proft, F. Reactivity of gas-phase, crystal and supported V2O5 systems studied using density functional theory based reactivity indices. Chem. Phys. Lett. 2008, 456, 59–63. [Google Scholar] [CrossRef] [Scilit]
- Biovia, D.S. BIOVIA Materials Studio, version 2017; Dassault Systèmes: San Diego, CA, USA, 2017.
- Ali, Z.; Wu, Y.-W.; Wu, Y.; Arain, Z.; Xu, M.-X.; Lu, Q.; Ma, H.; Zhao, H.-Y. Inhibition effects of Pb species on the V2O5-MoO3/TiO2 catalyst for selective catalytic reduction of NOx with NH3: A DFT supported experimental study. Appl. Surf. Sci. 2020, 525, 146582. [Google Scholar] [CrossRef] [Scilit]
- Scanlon, D.O.; Walsh, A.; Morgan, B.J.; Watson, G.W. An ab initio study of reduction of V2O5 through the formation of oxygen vacancies and Li intercalation. J. Phys. Chem. C 2008, 112, 9903–9911. [Google Scholar] [CrossRef] [Scilit]

Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2026 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license.