Skip to Content
  • 3.1
    Impact Factor
  • 5.9
    CiteScore
  • 14 days
    Submission to First Decision
  • 4 days
    Acceptance to Publication

Genes

Genes is a peer-reviewed, open access journal of genetics and genomics published monthly online by MDPI. The Epigenetics Society (ES) and the Spanish Society for Nitrogen Fixation (SEFIN) are affiliated with Genes, and their members receive discounts on the article processing charges.

Get Alerted

Add your email address to receive forthcoming issues of this journal.

All Articles (15,402)

  • Article
  • Open Access

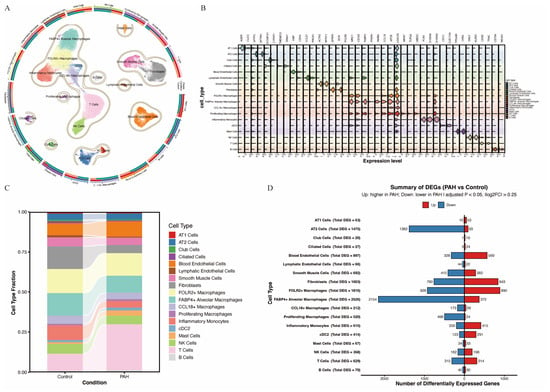
Background/Objectives: How macrophage-associated expression changes relate to fibroblast extracellular matrix (ECM) programs in pulmonary arterial hypertension (PAH) remains incompletely understood. This study aimed to characterize the relationship between clinically associated macrophage expression patterns and fibroblast ECM programs and to examine their spatial correspondence. Methods: We integrated FinnGen GWAS summary statistics with human single-cell and bulk lung transcriptomes. Candidate genes were evaluated using machine learning, hemodynamic associations, and cellular expression profiling. Cell communication and regulon inference were performed, followed by reanalysis of mouse spatial transcriptomic data comprising one wild-type section and one conditional knockout section. Results: The single-cell atlas comprised 55,158 cells, with fibroblasts among the populations showing the highest median trait relevance scores. An eight-gene tissue classifier achieved post-selection AUCs of 0.873 and 0.992 in two external cohorts used for model selection and evaluation. Bulk lung CD163 expression was inversely correlated with pulmonary vascular resistance (Spearman ρ = −0.522), while pooled single-cell summaries showed lower CD163 expression across four macrophage populations in PAH. The CD163–PVR association remained inverse but was attenuated after adjustment for clinical covariates and estimated macrophage fraction (p = 0.067). Fibroblast ECM-related pathway enrichment was also supported by sample-level analysis. CellChat predicted FN1- and SPP1-associated communication toward fibroblasts, although sample-level differences in macrophage FN1 and SPP1 expression and in fibroblast FN1 expression did not reach significance after multiple-testing correction (fibroblast FN1 FDR = 0.059). Among the candidate ECM-associated regulons, only ERG showed limited sample-level support (adjusted p = 0.049). Mouse spatial analysis showed shorter mean source-defined macrophage–fibroblast pair distances in the conditional knockout section, although the spatial block-bootstrap interval for the bin-level distance difference included zero. Within this section, detection of macrophage-associated Spp1 or Fn1 was associated with higher ECM scores in paired fibroblast-associated regions after excluding both ligand genes from the ECM signature. Conclusions: These analyses characterize co-occurring macrophage-associated expression changes, predicted macrophage–fibroblast communication, and fibroblast ECM programs in PAH. The mouse spatial reanalysis provides complementary spatial context for these observations, while the inferred communication and regulatory relationships should be considered computationally predicted and require further experimental validation.

Genes

20 November 2026

Integrated single-cell landscape and cell-type-specific transcriptional alterations in PAH. (A) UMAP visualization of the annotated cell populations, with accompanying cohort and condition information. (B) Stacked violin plots showing the expression of representative cell-type markers. (C) Pooled proportions of the annotated populations in control and PAH tissues. Connecting bands link the same cell populations between conditions and do not indicate cellular transitions. (D) Numbers of upregulated and downregulated genes in PAH relative to controls within each cell population. Differentially expressed genes were defined by an adjusted p value < 0.05 and an absolute average log2 fold change > 0.25.
  • Article
  • Open Access

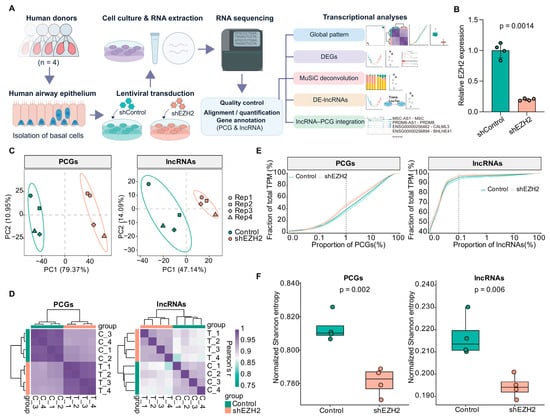
Background: Airway basal cells (BCs) maintain epithelial homeostasis and contribute to epithelial regeneration after injury. How enhancer of zeste homolog 2 (EZH2), a key epigenetic regulator, controls the transcriptional programs, especially the coding and non-coding transcriptional consequences underlying human airway basal cell states, remains poorly understood. Methods: Here, we performed rRNA-depleted total RNA sequencing of donor-matched control and shEZH2 primary human airway basal cells from four independent donors and integrated coding and non-coding transcriptomic analyses with basal-cell state inference and multi-criteria candidate prioritization. Results: EZH2 depletion broadly remodeled the BCs transcriptome. GSEA showed prominent negative enrichment of nucleosome organization, chromosome segregation, DNA replication, and cell-cycle programs, accompanied by positive enrichment of extracellular matrix- and signaling-related processes. Reference-guided analysis further revealed a marked reduction in the transcriptional similarity to the Cycling Basal reference state, accompanied by increased similarity to Basal and Suprabasal reference states, indicating a shift in the basal-cell transcriptional landscape away from a Cycling Basal-like program. In parallel, differentially expressed lncRNAs were associated with distinct transcriptional programs, while multi-criteria integration prioritized eight consensus lncRNA–PCG relationships for further investigation. Conclusions: These findings define coordinated coding and non-coding transcriptional remodeling following EZH2 knockdown in primary human airway basal cells and nominate candidate lncRNA–PCG regulatory relationships that may contribute to human airway basal-cell state regulation, providing a framework for understanding epigenetic regulation of basal-cell states and its potential relevance to airway epithelial remodeling in respiratory disease.

Genes

20 November 2026

Experimental design and global transcriptomic characteristics. (A) Schematic of the experimental design and analysis workflow, including primary airway basal cell culture, lentiviral transduction, RNA sequencing, and downstream transcriptomic analyses. (B) qPCR validation of EZH2 knockdown in four paired samples. Statistical significance was assessed using a paired two-tailed Student’s t-test. (C) PCA plots of PCG and lncRNA expression profiles across the eight samples. (D) Sample correlation heatmaps based on PCG and lncRNA expression profiles and the heatmap color scales indicate pairwise Pearson correlation coefficients (r). (E) Cumulative abundance distributions of PCGs and lncRNAs, with transcripts ranked by TPM. (F) Shannon entropy of PCG and lncRNA expression distributions. Statistical significance was assessed using a paired two-tailed Student’s t-test.
  • Article
  • Open Access

Background: The independent origins of plant parasitism in nematodes provide an ideal model for exploring the evolution of parasitic adaptations. Methods: Using a core set of 7 species and an expanded set of 16 species, we performed comparative genomic analyses across representative lineages of the suborder Tylenchina—specifically the superfamily Aphelenchoidea, family Anguinidae (superfamily Sphaerularioidea), and superfamily Tylenchoidea—to uncover the genetic basis of cross-superfamily molecular convergence and lineage-associated divergence. Results: Overall, sequence-level molecular convergence was relatively weak and may partly reflect the dilution of evolutionary signals by closely related ancestral backgrounds and the diverse genetic mechanisms driving the plant-parasitic phenotype. Nevertheless, independently evolved lineages exhibited recurrent functional themes in host interaction, barrier penetration, and nutrient acquisition. At the metabolic level, the unsaturated fatty acid biosynthesis pathway (ko01040) showed consistent contraction across all phylogenetic models, pointing to a potential shift toward reliance on host-derived lipids. At the sequence level, a subset of genes displayed site-level convergence validated through independent alignment sets; these include homologs implicated in host recognition, effector folding, nitrogen metabolism, and stress perception at the host–pathogen interface. Pathway counts formed a continuous gradient rather than distinct groups, with 31 pathways demonstrating significant contraction (BH-FDR corrected PGLMM-NB2 model; median β = −0.31 and −0.77). Notably, 7 of the 12 convergent genes mapped to pathways undergoing contraction—including lipid/sphingolipid metabolism, oxidative phosphorylation, protein homeostasis, and nitrogen metabolism—indicating that sequence-level convergence and pathway-level reduction converge on shared functional modules. Conclusions: Together, these findings establish a comprehensive comparative genomic framework for understanding the origin and evolutionary radiation of plant parasitism in nematodes.

Genes

20 November 2026

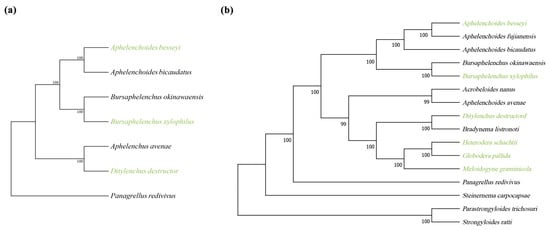
Maximum likelihood (ML) phylogenetic tree of nematodes constructed from single-copy orthologous genes. (a) Phylogenetic tree based on the seven core species (A. besseyi, A. bicaudatus, B. xylophilus, B. okinawaensis, D. destructor A. avenae and P. redivivus), with the free-living nematode P. redivivus placed outside the clade containing the six ingroup species; (b) expanded phylogenetic tree based on the 16 expanded-set species. Values on the nodes represent bootstrap support values from the maximum likelihood analysis (1000 replicates). Species names in green indicate plant-parasitic nematodes; those in black indicate non-plant-parasitic nematodes.
  • Article
  • Open Access

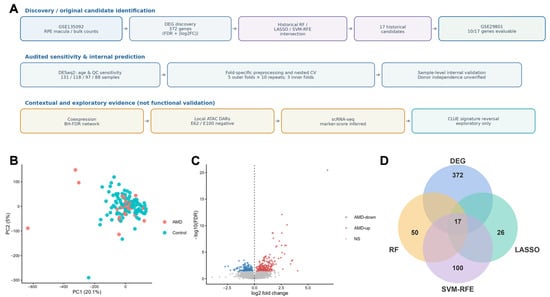
Background: Age-related macular degeneration (AMD) involves molecular changes in the retinal pigment epithelium (RPE) and choroidal tissues. Transcriptomic studies have identified candidate genes associated with AMD, although their reproducibility across cohorts and potential cellular and regulatory contexts require further investigation. Methods: We analyzed macular RPE/choroid RNA-seq data from GSE135092 (26 AMD and 105 control samples) and assessed candidate-gene expression in GSE29801 (79 AMD and 96 control samples). Six DESeq2 models evaluated age and QC sensitivity. Nested cross-validation assessed sample-level classification using training-fold-specific preprocessing and feature selection. Coexpression networks were assessed with target-wise Benjamini–Hochberg adjustment. We also examined single-cell expression, ATAC-seq accessibility, and HiChIP contacts. Results: The original discovery analysis identified 372 genes with FDR < 0.05 and |log2FC| > 0.5. Ten of 17 candidate genes were evaluable in GSE29801; seven lacked usable GPL4133 probes. PRSS33 and CHIT1 showed positive AMD-related expression effects in GSE29801. In GSE135092, PRSS33 remained directionally increased across sensitivity models but lost FDR significance after QC exclusion and in age-complete subsets. Nested cross-validation achieved mean repeat-level ROC-AUCs of 0.714, 0.699, and 0.760 for random forest, logistic LASSO, and SVM-RFE, respectively. PRSS33 was detected in 0.528% of marker-score-inferred RPE cells. E100 and E62 showed decreased ATAC-seq accessibility in AMD. Conclusions: PRSS33 showed directionally concordant expression changes across two AMD RPE/choroid cohorts and remains an exploratory candidate. Its association in the discovery cohort was sensitive to QC exclusion and age-complete analyses. The available single-cell and chromatin data provide preliminary biological context; donor-level and functional studies are needed to clarify its cellular distribution and potential regulatory relationships.

Genes

20 November 2026

Study overview and candidate discovery. (A) Discovery in GSE135092, external expression assessment in GSE29801, and sensitivity and internal modeling analyses. (B) PCA of the 131 discovery samples. (C) Volcano plot based on the discovery thresholds; the dashed vertical line indicates log2 fold change = 0. (D) Original overlap of DEGs and three machine-learning selection methods, yielding 17 candidates.

Highly Accessed Articles

News & Conferences

Latest Issues

Open for Submission

Journal Sections

XFacebookLinkedIn
Genes - ISSN 2073-4425