


Abstract
Neonatal calf diarrhea (NCD) is one of the most important problems of calf breeding across the world. It causes deaths in calves in the first 10 days of their life, and it is mainly caused by Escherichia coli(E. coli), Bovine Rotavirus (BRV) and Bovine Coronavirus (BCoV). The lack of vaccines with consistently high protective efficacy against the main causes of NCD makes disease control highly challenging. The current study aims to design a multi-epitope mRNA-based vaccine targeting the major pathogens responsible for NCD using immunoinformatic tools and molecular modeling approaches. BRV capsid protein VP6, BCoV Spike glycoprotein and E. coli F5 fimbrial protein were used as antigenic proteins to predict potential epitopes. Fifteen selected epitopes were linked with suitable linkers and conjugated with a built-in adjuvant, resulting in the design of a stable, antigenic and non-allergenic vaccine candidate against NCD pathogens. Furthermore, molecular docking analysis shows strong binding affinity between the vaccine candidate and the bovine toll-like receptors TLR2 and TLR4 at low energy and high stability. Based on these findings, the proposed multi-epitope vaccine represents a promising approach for the prevention and control of neonatal calf diarrhea and provides a solid scientific foundation for future experimental studies to validate its efficacy and safety in vivo.
1. Introduction
Neonatal calf diarrhea (NCD) is one of the most significant health problems affecting newborn calves and represents a major cause of morbidity and mortality during the first weeks of life. While several pathogens are responsible for NCD in newborn calves, Bovine Coronavirus (BCoV), Bovine Rotavirus (BRV) and Escherichia coli K99 are the main and economically important causative agents of the disease [1,2].
BRV and BCoV are viral causes of NCD. In the case of BRV, the virus infects mature enterocytes that are located at the tips of the small intestine. This leads to impaired digestion and absorption, resulting in diarrhea, dehydration, electrolyte imbalance and growth retardation. In the same manner, BCoV infects the small and large intestine, causing severe and prolonged diarrhea. Several studies have documented increased severity and duration of diarrhea in the case of co infection with BCoV and BRV as compared with rotavirus infections alone [3]. In contrast to BCoV and BRV, E. coli K99 does not invade the intestinal epithelium but colonizes the gut through the K99 antigen, which binds to specific receptors on the intestinal epithelial cells. Then, heat-stable enterotoxins produced by the bacteria induce diarrhea due to stimulation, causing the excessive secretion of fluids and electrolytes into the intestinal lumen [1].
Several studies have documented that mixed infections involving viral and bacterial pathogens are frequently reported in neonatal calves and are associated with more severe clinical signs. These signs include high mortality rates and reduced response to treatment compared to single-pathogen infections. The interaction between Bovine BCoV, BRV and E coli K99 exacerbates intestinal damage and disrupts normal immune responses, increasing disease severity and economic losses in the cattle industry [2].
Currently, live-attenuated and inactivated vaccines are the main types of vaccines commercially used against the principal causes of NCD. However, these vaccines have several limitations, including incomplete coverage of all major causative agents, potential safety concerns and limited suitability for administration during the late stage of pregnancy [3]. NCD remains a serious problem for livestock farms due to a lack of effective vaccines that can provide protective immunity for calves against causative agents. Therefore, development of a vaccine candidate targeting multi-pathogens of the disease is urgently needed. In recent years, advances in molecular biology and immunoinformatic have facilitated the design of next-generation vaccines, including multi-epitope and mRNA-based vaccines, which offer several advantages over traditional vaccine platforms [4].
Recently, reverse vaccinology was used to design and evaluate vaccines against a wide range of infectious pathogens. Most of these studies suggest that multi-epitope vaccines are able to stimulate both humoral and cellular responses. In this type of vaccine, epitopes from different pathogens can be included in a single vaccine, which provides a broad spectrum of protection. Furthermore, the combination these types of vaccines with the mRNA platform provides a high level of safety, rapid production and strong immune response without the risk of genomic integration [5,6,7]. The development in molecular technique facilities the constructing process of a multi-epitope vaccine and enables the incorporation of genetic elements such as built-in adjuvants, untranslated regions (UTRs), Kozak sequences and strong promoters in vaccine construct, thereby enhancing antigen expression and immune stimulation, further improving vaccine efficacy [8].
Considering the high prevalence of mixed infections caused by Bovine Coronavirus, Bovine Rotavirus and Escherichia coli K99, the development of a heterogeneous multi-epitope mRNA vaccine represents a promising approach for the prevention and control of neonatal calf diarrhea. As such, this vaccine strategy has the potential to provide broad, effective and long-lasting immunity, ultimately reducing disease burden and economic losses in the cattle industry. Our proposed vaccine is designed to target three major causative agents within a single vaccine and is intended for safe administration to dams during the final stage of pregnancy, thereby enhancing protective immunity in newborn calves.
2. Materials and Methods
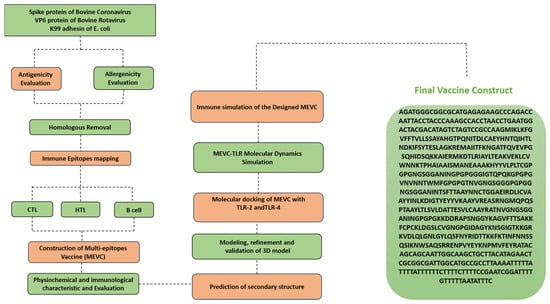
Figure 1 represents the workflow of the current study including design, prediction and immunoinformatic evaluation processes of the proposed vaccine.
Figure 1.
The flow chart represents the workflow of the study. The chart shows the processes for the design of MEVC, starting from choosing the antigenic proteins, immune simulation and then construction of the final form of the vaccine.
2.1. Sequence Retrieval of Protein
Protein sequences of the target antigens from Bovine Coronavirus (BCoV), Bovine Rotavirus (BRV) and Enterotoxigenic Escherichia coli (ETEC) K99 (F5) were retrieved from the National Center for Biotechnology Information (NCBI) database. For BCoV, the spike (S) protein was selected due to its crucial role in viral attachment and induction of neutralizing antibodies. For BRV, the inner capsid/middle layer protein VP6 was chosen as a major immunogenic antigen involved in protective immunity. For ETEC, the fimbrial protein K99 (F5) that mediates bacterial adhesion to the intestinal epithelial cells in neonatal calves was selected as a key virulence and vaccine target antigen. These proteins were selected based on their reported immunogenicity and relevance in the pathogenesis of neonatal calf diarrhea [1,9].
2.2. Epitope Mapping
2.2.1. Epitope Prediction Tools and Parameters
All epitope predictions were performed using the Immune Epitope Database (IEDB) Analysis Resource (http://tools.iedb.org). Default recommended settings were applied for all predictions, as these represent IEDB’s optimized parameters for each prediction type [10,11].
MHC Class I (CTL) epitopes: Predictions were performed using the IEDB “recommended” method, which automatically selects the consensus method (combining ANN, SMM, and Comblib) where available and falls back to NetMHCpan-4.1 for other allele-length combinations. NetMHCpan-4.1 has demonstrated high predictive accuracy for bovine MHC class I molecules.
MHC Class II (HTL) epitopes: The IEDB “recommended” method was used, comprising the consensus method (which integrates NN-align, SMM-align, and combinatorial peptide scanning library) as the primary approach, with NetMHCIIpan as a fallback.
Linear B cell (LBL) epitopes: The BepiPred-2.0 algorithm was employed, which uses a random forest classifier trained on epitope and non-epitope residues from crystal structures.
2.2.2. Bovine MHC (BoLA) Alleles Selected
The BoLA alleles were used for epitope prediction, selected based on published frequency data in Holstein cattle (the predominant dairy breed worldwide) [12].
2.3. Bovine Homology
BLASTp (https://blast.ncbi.nlm.nih.gov, accessed on: 12 February 2025) was used to analyze of the predicted epitopes against the cattle proteome (TaxID: 9913) in the NCBI Database and also to identify any potential cross reactions. The server’s default settings were used. Only peptides with E-value > 0.05 were retained to avoid self-reactivity.
2.4. Assessment of B Cell and T Cell Epitopes
Antigenicity of the predicted epitopes derived from the Immune Epitope Database (IEDB) analysis was evaluated using the VaxiJen v2.0 web server, with a threshold value of 0.5 to identify highly antigenic epitopes [13]. Toxicity of the predicted epitopes was estimated using ToxinPred 3.0 Web Server (https://webs.iiitd.edu.in/raghava/toxinpred/ (accessed on: 12 February 2025)). Only epitopes exceeding the defined threshold were considered antigenic and selected for further analysis. Subsequently, the allergenicity of the shortlisted epitopes was assessed using the AllerTOP v2.0 server (https://www.ddg-pharmfac.net/AllerTOP/ (accessed on: 12 February 2025)), which predicts the allergenic potential based on physicochemical properties [14]. Epitopes predicted as non-allergenic and antigenic were retained for construction of the multi-epitope vaccine candidate.
2.5. Assembly of the Final Multi-Epitope mRNA Vaccine
To construct the final multi-epitope mRNA vaccine candidate, highly conserved, antigenic, non-allergenic and overlapping epitopes derived from Bovine Coronavirus, Bovine Rotavirus and Escherichia coli K99 were assembled into a single vaccine construct. The selected epitopes were joined using appropriate linkers, including GPGPG, AAY and KK to maintain the independent immunogenicity of each epitope and to prevent junctional immunogenicity [15]. A built-in adjuvant sequence was incorporated into the vaccine construct to enhance immune stimulation. Additionally, molecular optimization elements such as optimized 5′ and 3′ untranslated regions (UTRs) and a Kozak sequence were included to ensure efficient antigen expression and translation. This rational design strategy aims to enhance the stability, immunogenicity and overall efficacy of the proposed heterogeneous multi-epitope mRNA vaccine.
The complete optimized nucleotide sequence and the corresponding amino acid sequence of the proposed vaccine construct are provided in Supplementary Materials.
2.6. Profile of Physicochemical and Immunological Properties of the Construct
The physicochemical properties of the designed multi-epitope vaccine construct were evaluated using the ProtParam tool available on the ExPASy server (https://web.expasy.org/protparam/ (accessed on: 13 February 2025)) [16]. The analyzed parameters included molecular weight, amino acid composition, theoretical isoelectric point (pI), aliphatic index, estimated half-life in vitro and in vivo based on the N-end rule, and the grand average of hydropathicity (GRAVY) value.
The antigenicity of the vaccine construct was predicted using the VaxiJen v2.0 web server, with a threshold value of 0.5 to identify the high antigenic candidates [13]. Furthermore, the allergenicity of the construct was assessed using the AllerTOP v2.0 server to ensure the safety of the designed vaccine by predicting its non-allergenic nature [14].
2.7. Prediction of Secondary Structure (2D)
The secondary structural features of the vaccine construct, including alpha-helices, beta-turns, and random coils were predicted using the PSIPRED v3.3 web server [17]. This analysis provided insight into the structural stability and folding patterns of the designed vaccine.
2.8. Prediction of Tertiary Structure (3D)
The three-dimensional (tertiary) structure of the constructed vaccine was predicted using the I-TASSER web server (https://zhanggroup.org/I-TASSER/ (accessed on: 16 February 2025)) [18]. Among the generated models, the best-quality 3D structure was selected based on the confidence score (C-score), which typically ranges from −5 to +2, where a higher score indicates greater reliability of the predicted model.
2.9. Validation and Refinement of the Vaccine Model
The three-dimensional structure (3D) of the designed multi-epitope vaccine was refined using the GalaxyRefine web server (https://galaxy.seoklab.org/refine (accessed on: 16 February 2025)) to improve the overall structural quality and stability of the model [19]. The refinement process enhanced side-chain conformations and reduced structural clashes, resulting in a more reliable vaccine model. The refined vaccine structure was then validated using the RAMPAGE web server (https://bio.tools/RAMPAGE, accessed on: 12 February 2025) to analyze the Ramachandran plot, which assesses the stereochemical quality of the protein model [20]. The majority of amino acid residues were found within the favored and allowed regions, indicating good backbone geometry and acceptable conformational accuracy of the modeled vaccine structure. These validation results confirm the structural reliability and suitability of the refined vaccine model for subsequent immunoinformatic analyses, including molecular docking and immune interaction studies.
2.10. Molecular Docking of the Vaccine Candidate with Immune Receptors
Molecular docking analysis was performed to evaluate the interaction between the designed multi-epitope vaccine and TLR-2 and TLR-4 receptors using the ClusPro v2.0 web server (https://cluspro.bu.edu) [21].
The three-dimensional structures of TLR-2 and TLR-4 were retrieved from the Protein Data Bank (PDB) and prepared for docking analysis. The modeled vaccine construct was docked separately with each receptor to assess binding affinity and interaction stability.
The best docked complexes were selected based on the lowest binding energy scores and optimal cluster size. Visualization and structural analysis of the docked complexes were performed using PyMOL 3.1 to identify key binding residues and interaction interfaces between the vaccine construct and both TLR-2 and TLR-4 receptors. Additionally, the HDOCK v2.5 web server was employed to further validate the docking results and enhance the reliability of the predicted interactions. These analyses suggest the designed vaccine might stimulate immune responses through effective interaction with TLR-2 and TLR-4 receptors.
2.11. Normal Mode Analysis (NMA)
Normal mode analysis (NMA) was performed using the iMODS server ((http://imods.chaconlab.org) to evaluate the intrinsic flexibility and collective motions of the receptor–vaccine complex. The complexes with the most favorable energy have been selected for the analysis. Through the iMODS server, the stability with molecules and atoms was assessed.
2.12. Immune Simulation of the Vaccine Candidate
The C-ImmSim server v6 [22] was used to assess the ability of the proposed vaccine to stimulate a protective immune response within the host against the target pathogens. The proposed vaccine was evaluated based on three injections at four-week intervals.
3. Results
3.1. Retrieval and Analysis of Selected Proteins Sequences
Bovine Coronavirus spike protein (BCoV, Accession no. P15777); Bovine Rotavirus VP6 protein (BRV, Accession no. Q68YC9) and Escherichia coli K99 fimbrial protein (K99, Accession no. WP_089508147.1) were selected as the target antigens. All three proteins were analyzed for antigenicity using the VaxiJen v2.0 web server, and the results indicated that all proteins are antigenic and capable of eliciting immune responses, with scores exceeding the threshold values.
3.2. Prediction and Evaluation of T and B Cell Epitopes
All potential epitopes in the three proteins were screened to select fully conserved, antigenic, and non-allergenic epitopes. From the Bovine Coronavirus spike protein (BCoV spike protein): 2 THL epitopes, 2 CTL epitopes and 2 LBL epitopes were selected. While 2 THL epitopes, 1 CTL epitope and 1 LBL epitope were selected from the Bovine Rotavirus VP6 protein (BRV VP6 protein). Finally, 2 THL epitopes, 2 CTL epitopes and 1 LBL epitope were selected from Escherichia coli K99 protein (E. coli K99 protein). As shown in Table 1, Table 2 and Table 3, all selected epitopes were combined to construct the proposed vaccine. The bovine BoLA reference set was operated for this analysis. Epitope selection was based on representative BoLA alleles available in IEDB rather than breed-specific allele frequency data.
Table 1.
Final selected CTL epitopes for the candidate vaccine.
Table 2.
Final selected HTL epitopes for the candidate vaccine.
Table 3.
Final selected LBL epitopes for the candidate vaccine.
3.3. Construction of the Multi-Epitope mRNA Vaccine
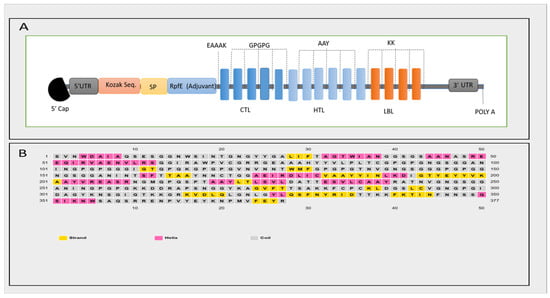
As show in Figure 2A, five CTL, six HTL and four LBL epitopes were selected to construct the proposed vaccine. AAY, GPGPG, and KK linkers were employed to fuse the epitopes and to prevent the epitope from interfering. To enhance immunogenicity, RpfE was incorporated as a built-in adjuvant, along with the Kozak sequence at the 5′ end via the EAAAK linker. RpfE from Mycobacterium tuberculosis H37Rv (NCBI NP_216966.1; UniProt O53177) is described as a TLR4 agonist that induces Th1/Th17 responses.
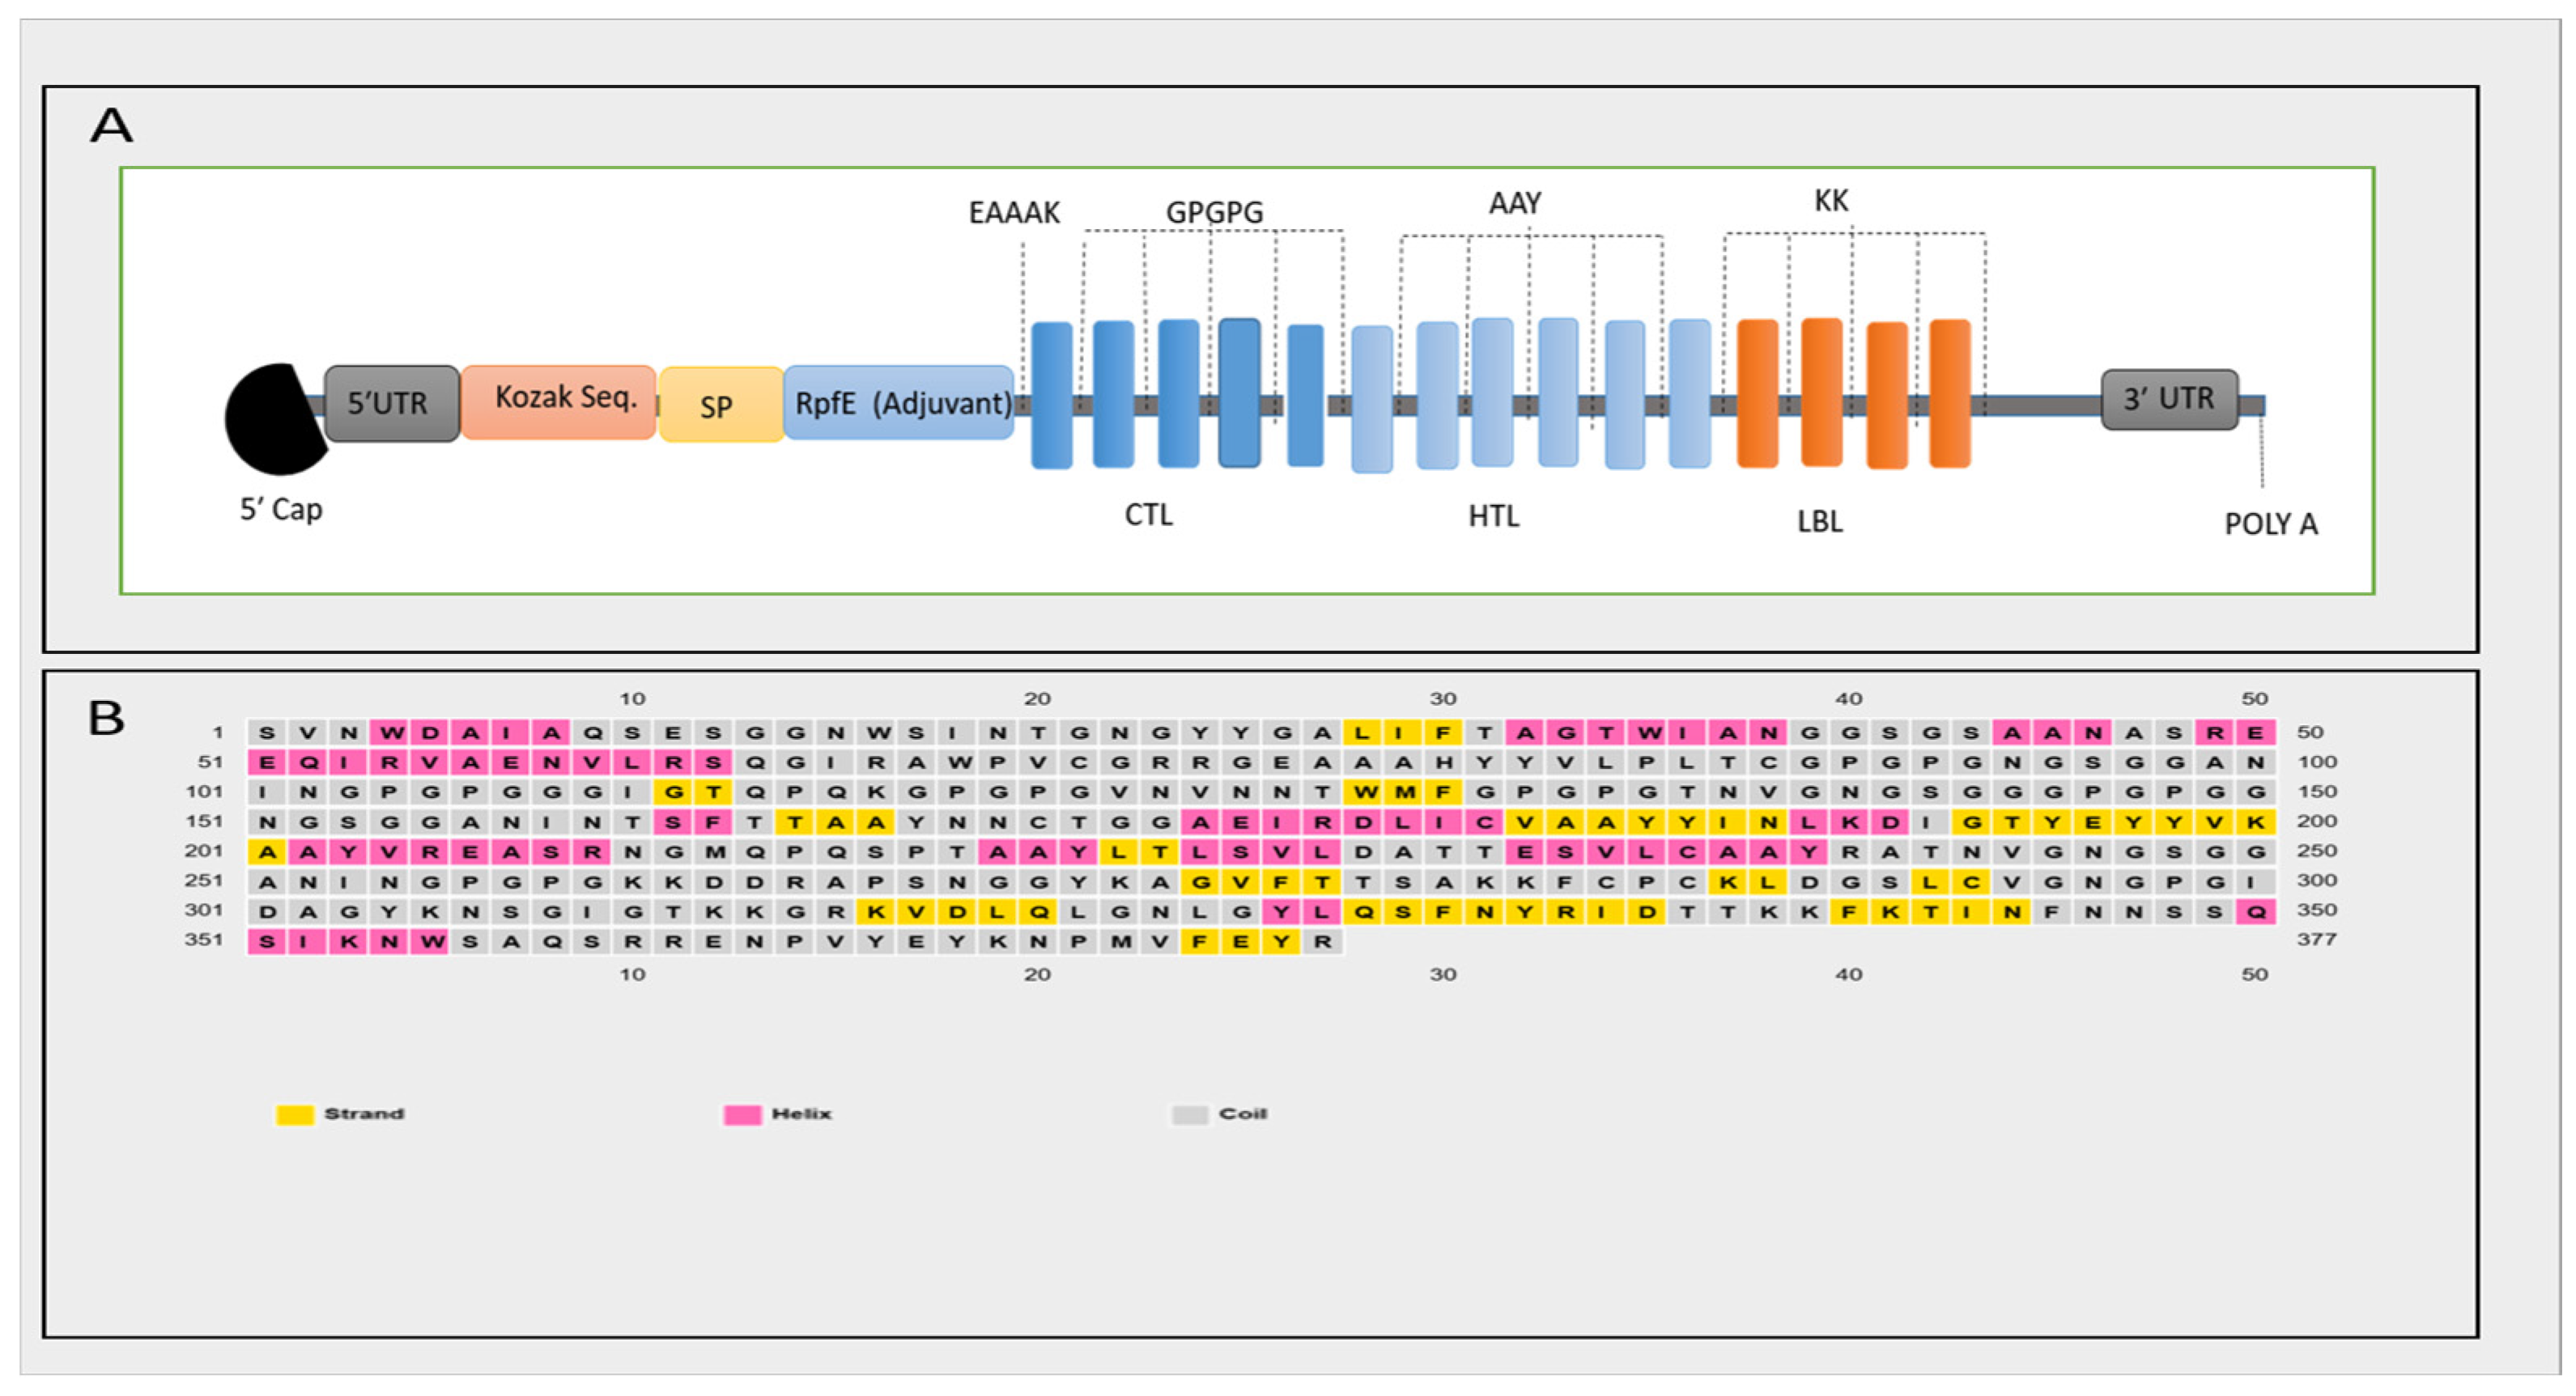
Figure 2.
Graphical map and secondary structure of the final mRNA multi-epitope vaccine construct. (A): The construct includes Kozak sequence and RpfE adjuvant at the N-terminal linked by EAAAK. HTL epitopes from BCoV, BRV and E. coli were joined using GPGPG linker, CTL epitopes using AAY linker and B-cell epitopes using KK linker. The 5′ and 3′ UTRs and Poly(A) tail were added to enhance mRNA stability and expression. (B) Secondary structure of the vaccine construct. The structure composite of 20.7% (78/377) α-helices, 13% (49/377) extended strands, and 66.3% (250/377) random coils.
Additionally, 5′ and 3′ untranslated regions (UTRs) were added to ensure stability and optimal expression.
3.4. Physicochemical and Immunological Analysis of the Proposed Vaccine
Immunogenicity of the vaccine was primarily predicted using VaxiJen and AllerTOP v1.1. The result shows that the proposed vaccine is antigenic with an antigenic score two-times more than the threshold score while probably not being an allergen to the selected host. The ProtParam web server was used to predict the physicochemical properties of the proposed vaccine. Table 4 summarizes the main physicochemical characters of the proposed vaccine.
Table 4.
Immunogenic and physicochemical characteristics of the vaccine construct.
3.5. Secondary and 3D Structure of the Proposed Vaccine Construct
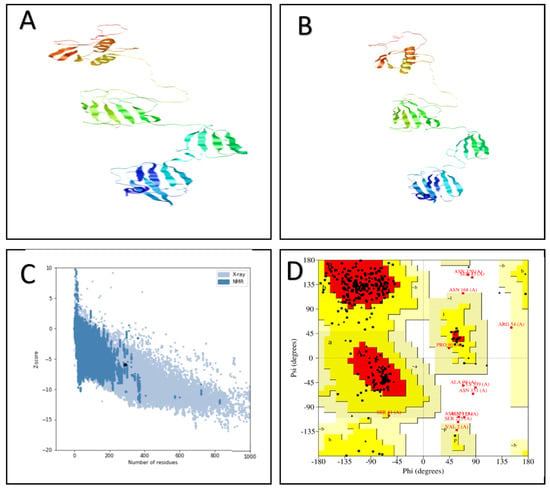
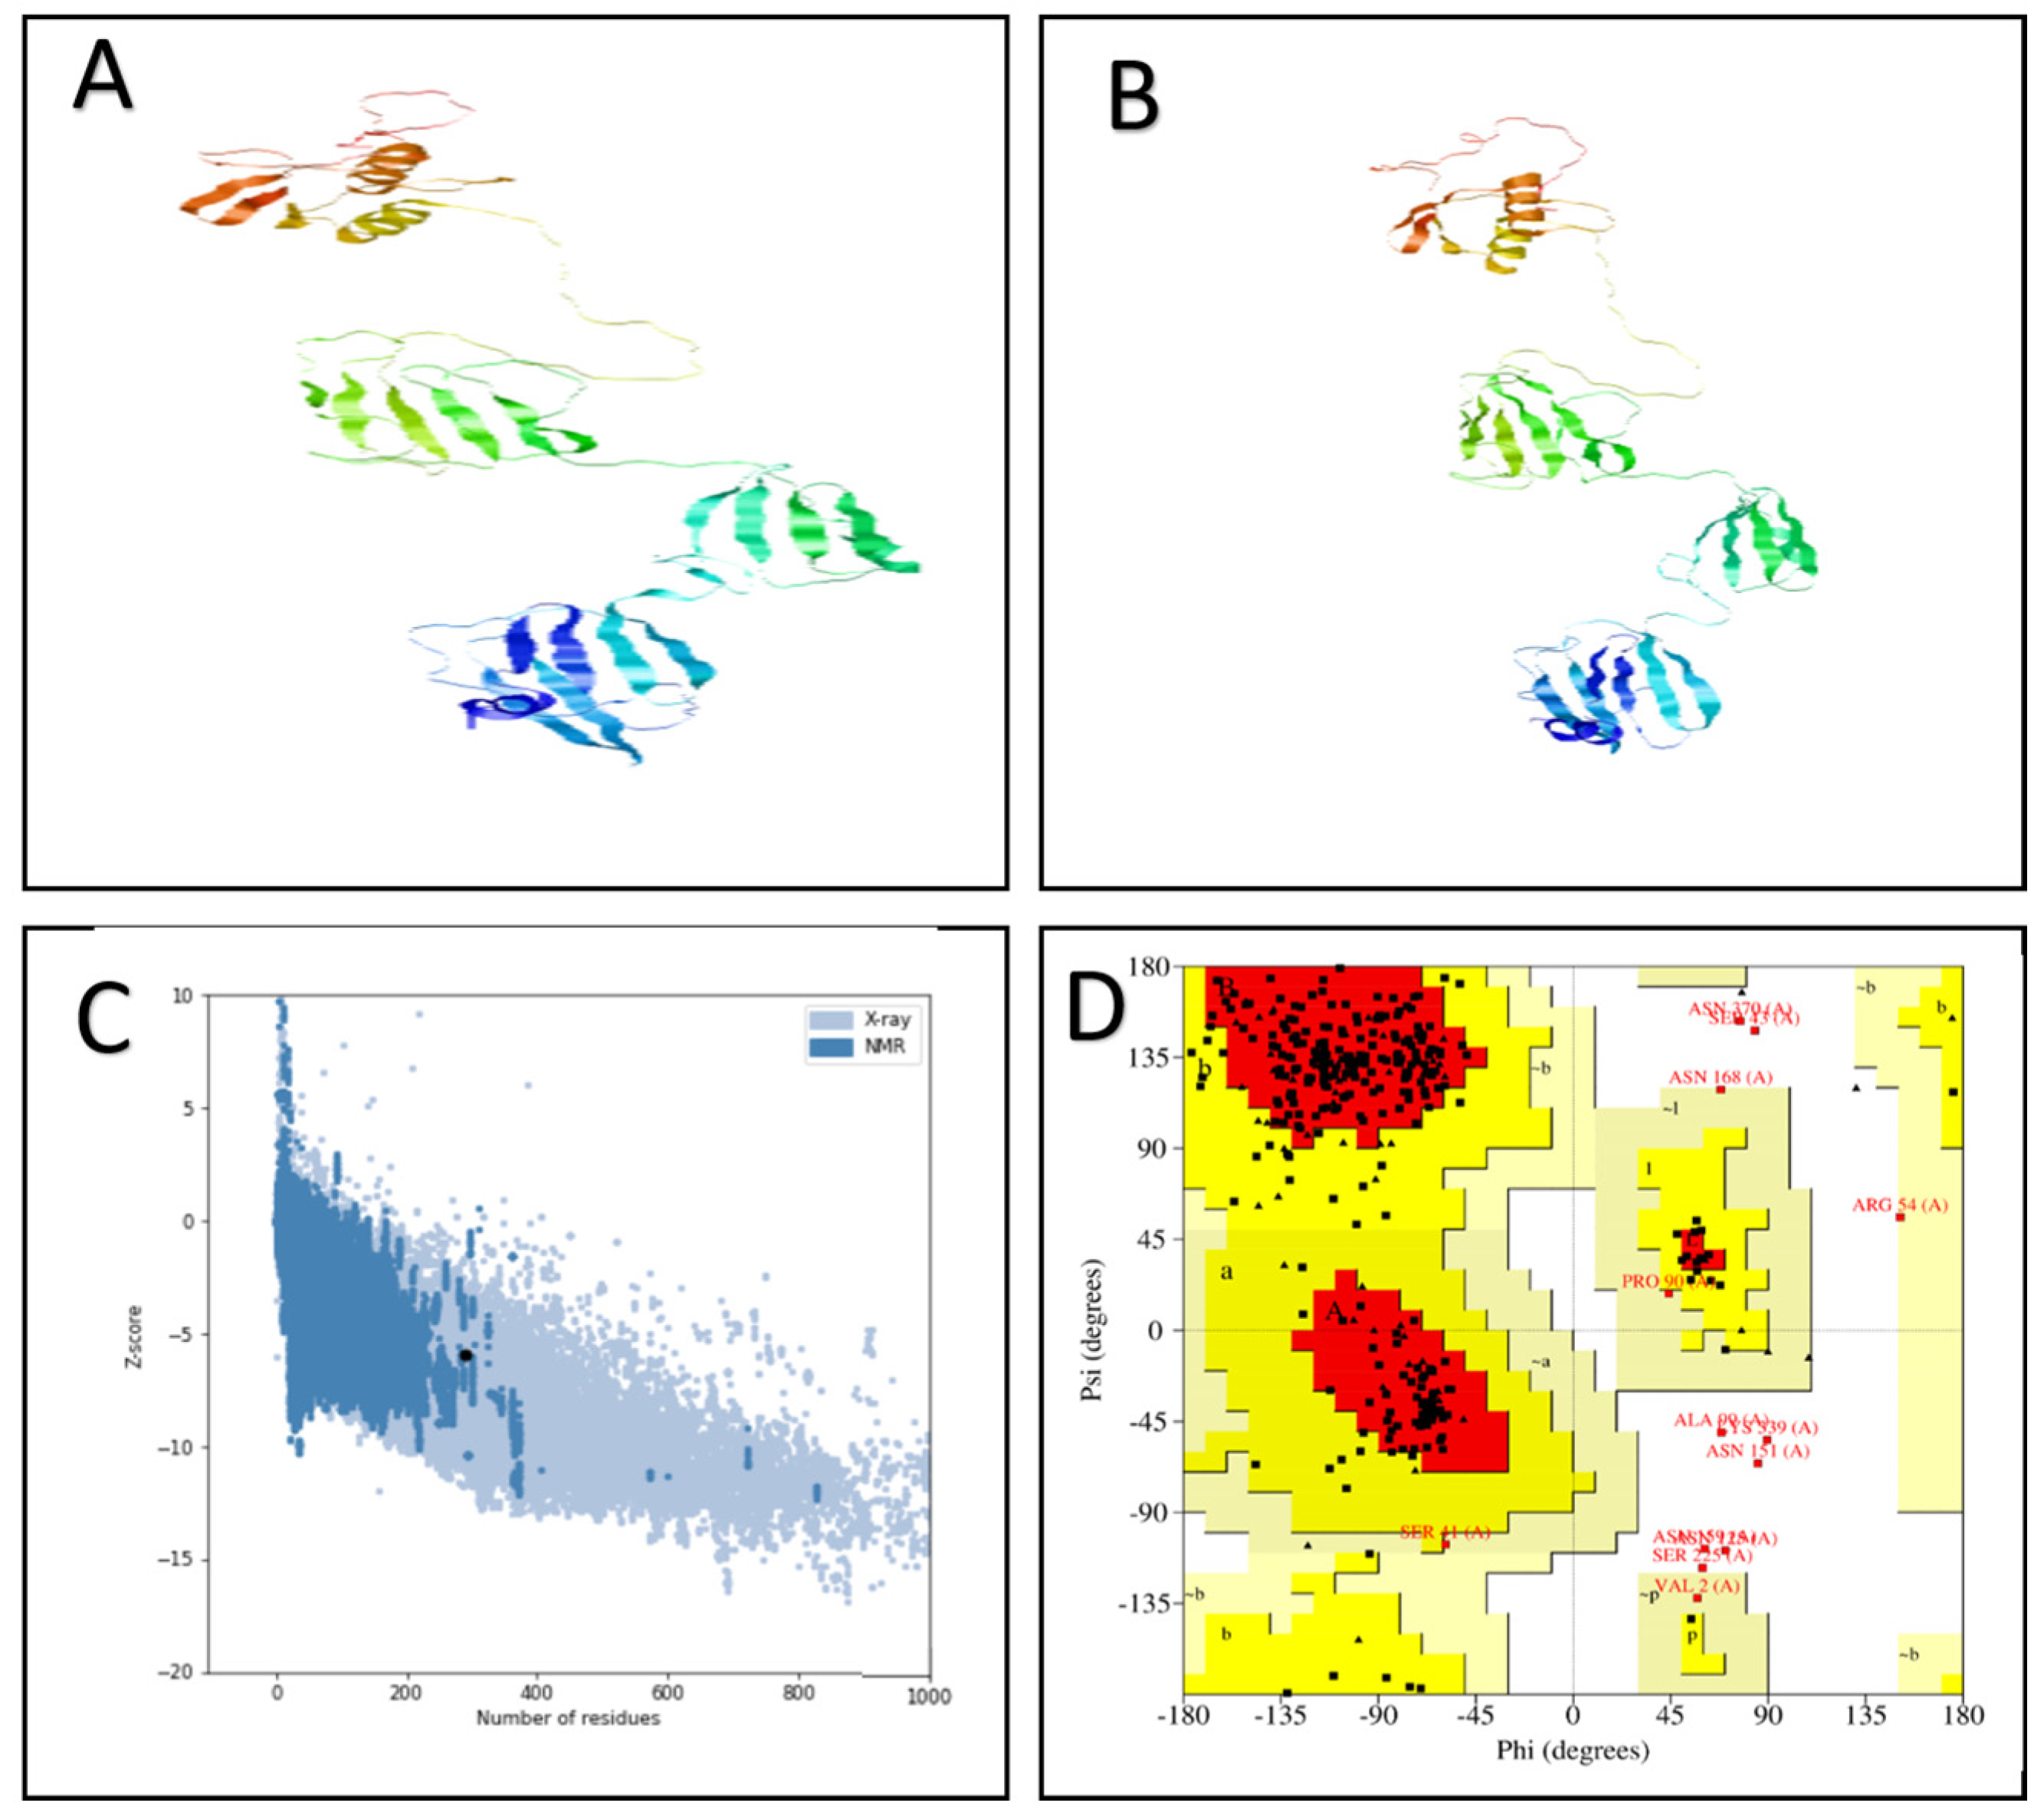
Figure 2B shows that the vaccine contained 20.7% (78/377) α-helices, 13% (49/377) extended strands, and 66.3% (250/377) random coils. The predicted secondary structure of the proposed vaccine refers to the high possibility of being able to recognize it by immune cell. A 3D structure of the vaccine construct was predicted using the I-TASSER server. The server predicts five models (1–5) with different com Scores (CS) of −2.15, −2.32, −3.14, −3.66 and −3.66, respectively. We chose model one with a CS of −2.15 (Figure 3A) then the selected model was refined with TheGalaxy Refine server. The model in (Figure 3B) was selected among five refined models based on five structural validation metrics, including GDT-HA score, RMSD score, MolProbity score, and Clash score. The selected models recorded a GDT-HA score of 0.8879, RMSD score of 0.571, MolProbity score of 2.346 and Clash score of 15.7, which is the best validation score among the five predicted models. As shown in Figure 3C, the Z-score of the refined model was −2.88 (Figure 3C). Other validation and quality scores of the refined structure were predicted using the SAVE 6.1 web server. The structure has an overall quality factor of 69.8925, while the Ramachandran plot (Figure 3D) shows that 97.2% of residues were in allowed regions and 0.9% were in disallowed regions. Overall, the predicted validation metrics represent a good quality of model and could be used for further analysis.
Figure 3.
Prediction, refinement and validity of the proposed vaccine construct. (A): Initial predicted model. (B): Refine of the vaccine structure with the galaxy web server. (C): Z-score calculated with the Pro-SA web server. (D): Ramachandran plot analysis. The red and yellow shaded regions represent the most favored and additionally allowed conformational regions, respectively, while the white areas correspond to disallowed regions. Black squares (■) indicate non-glycine residues and black triangles (▲) indicate glycine residues. The capital letters (A, B, L) denote the major favored conformational regions corresponding to right-handed α-helices, β-sheets, and left-handed α-helices, respectively, whereas the lowercase letters (a, b, l, p) indicate additionally allowed regions for α-helices, β-sheets, left-handed α-helices, and polyproline II helices, respectively.
3.6. Prediction of Discontinuous B-Cell Epitopes
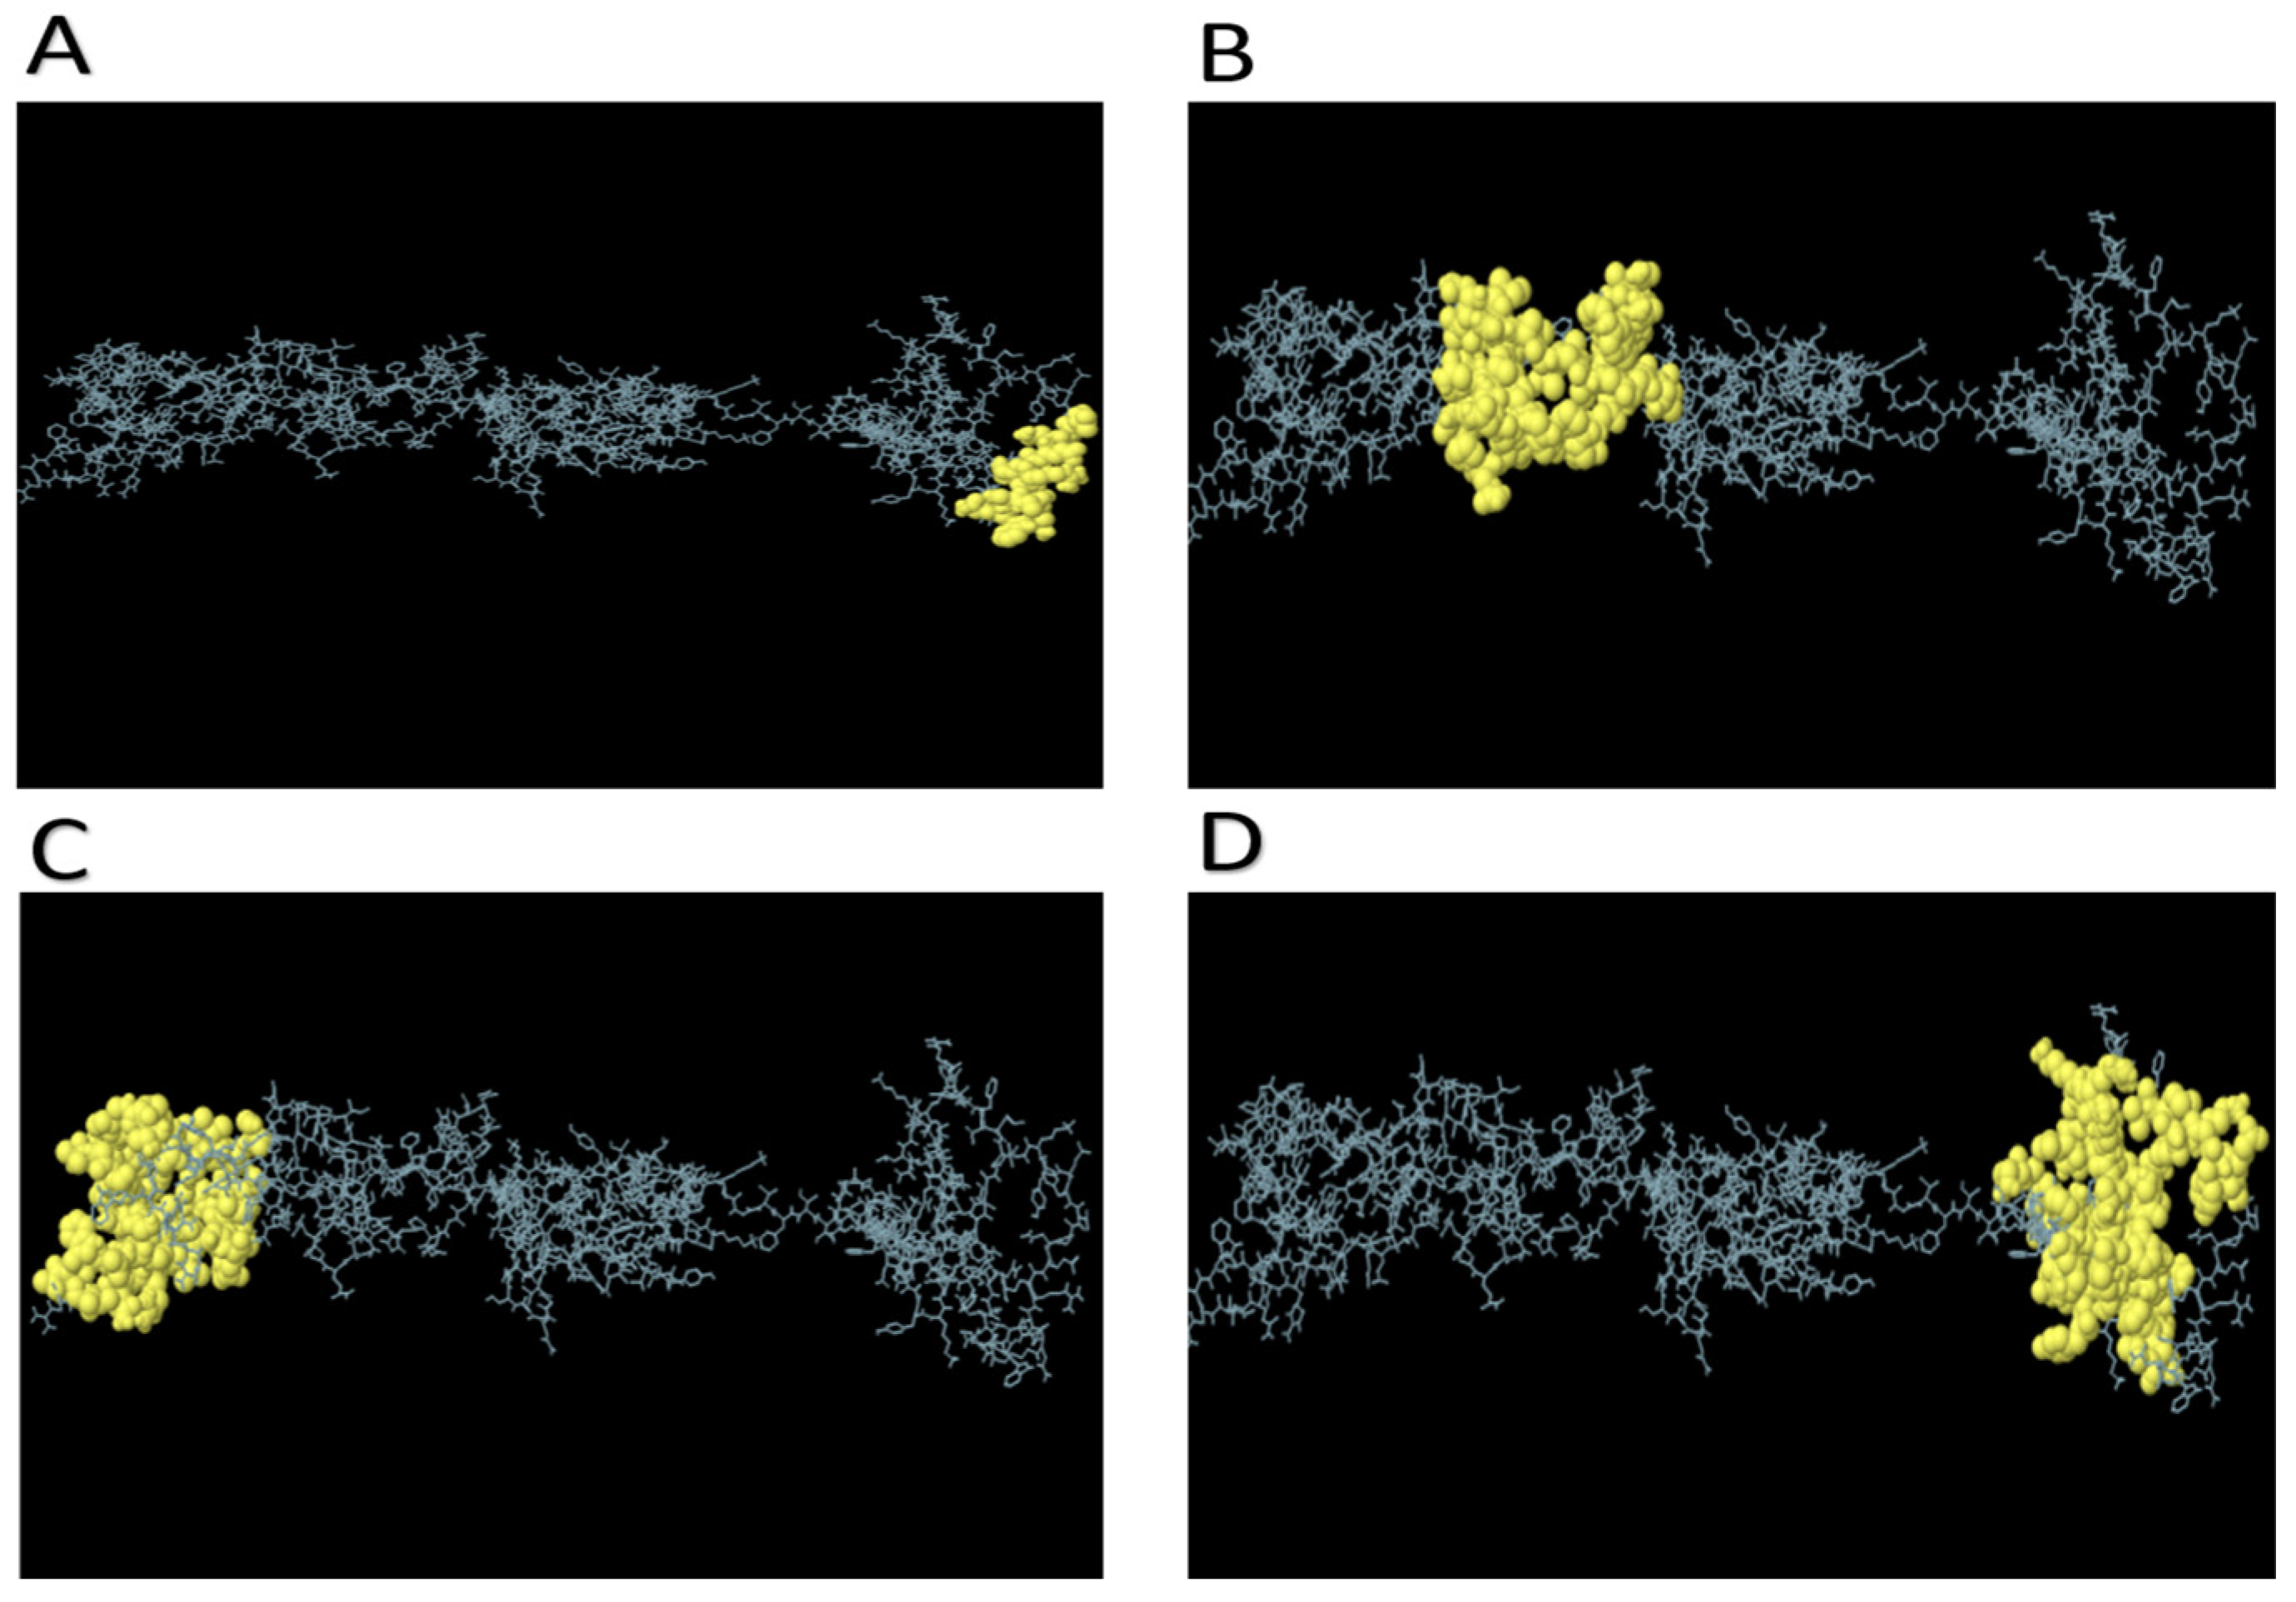
Analysis of the proposed vaccine with the ElliPro 1.0 server shows that this vaccine contains four discontinuous B-Cell epitopes (Figure 4). All predicted epitopes have scores above 0.5, with different numbers of amino acids. Table 5 shows the predicted epitopes and number of residues along with score for each predicted epitope.
Figure 4.
The 3D model predicted by the ElliPro server displaying discontinuous B-cell epitopes in response to the vaccine candidate. (A): Epitope with 12 residues. (B): Epitope with 48 residues. (C): Epitope with 63 residues. (D): Epitope with 62 residues.
Table 5.
The discontinuous B cell epitopes residues of the vaccine candidate.
3.7. Molecular Docking of the Vaccine Candidate with Bovine TLR2 and TLR4
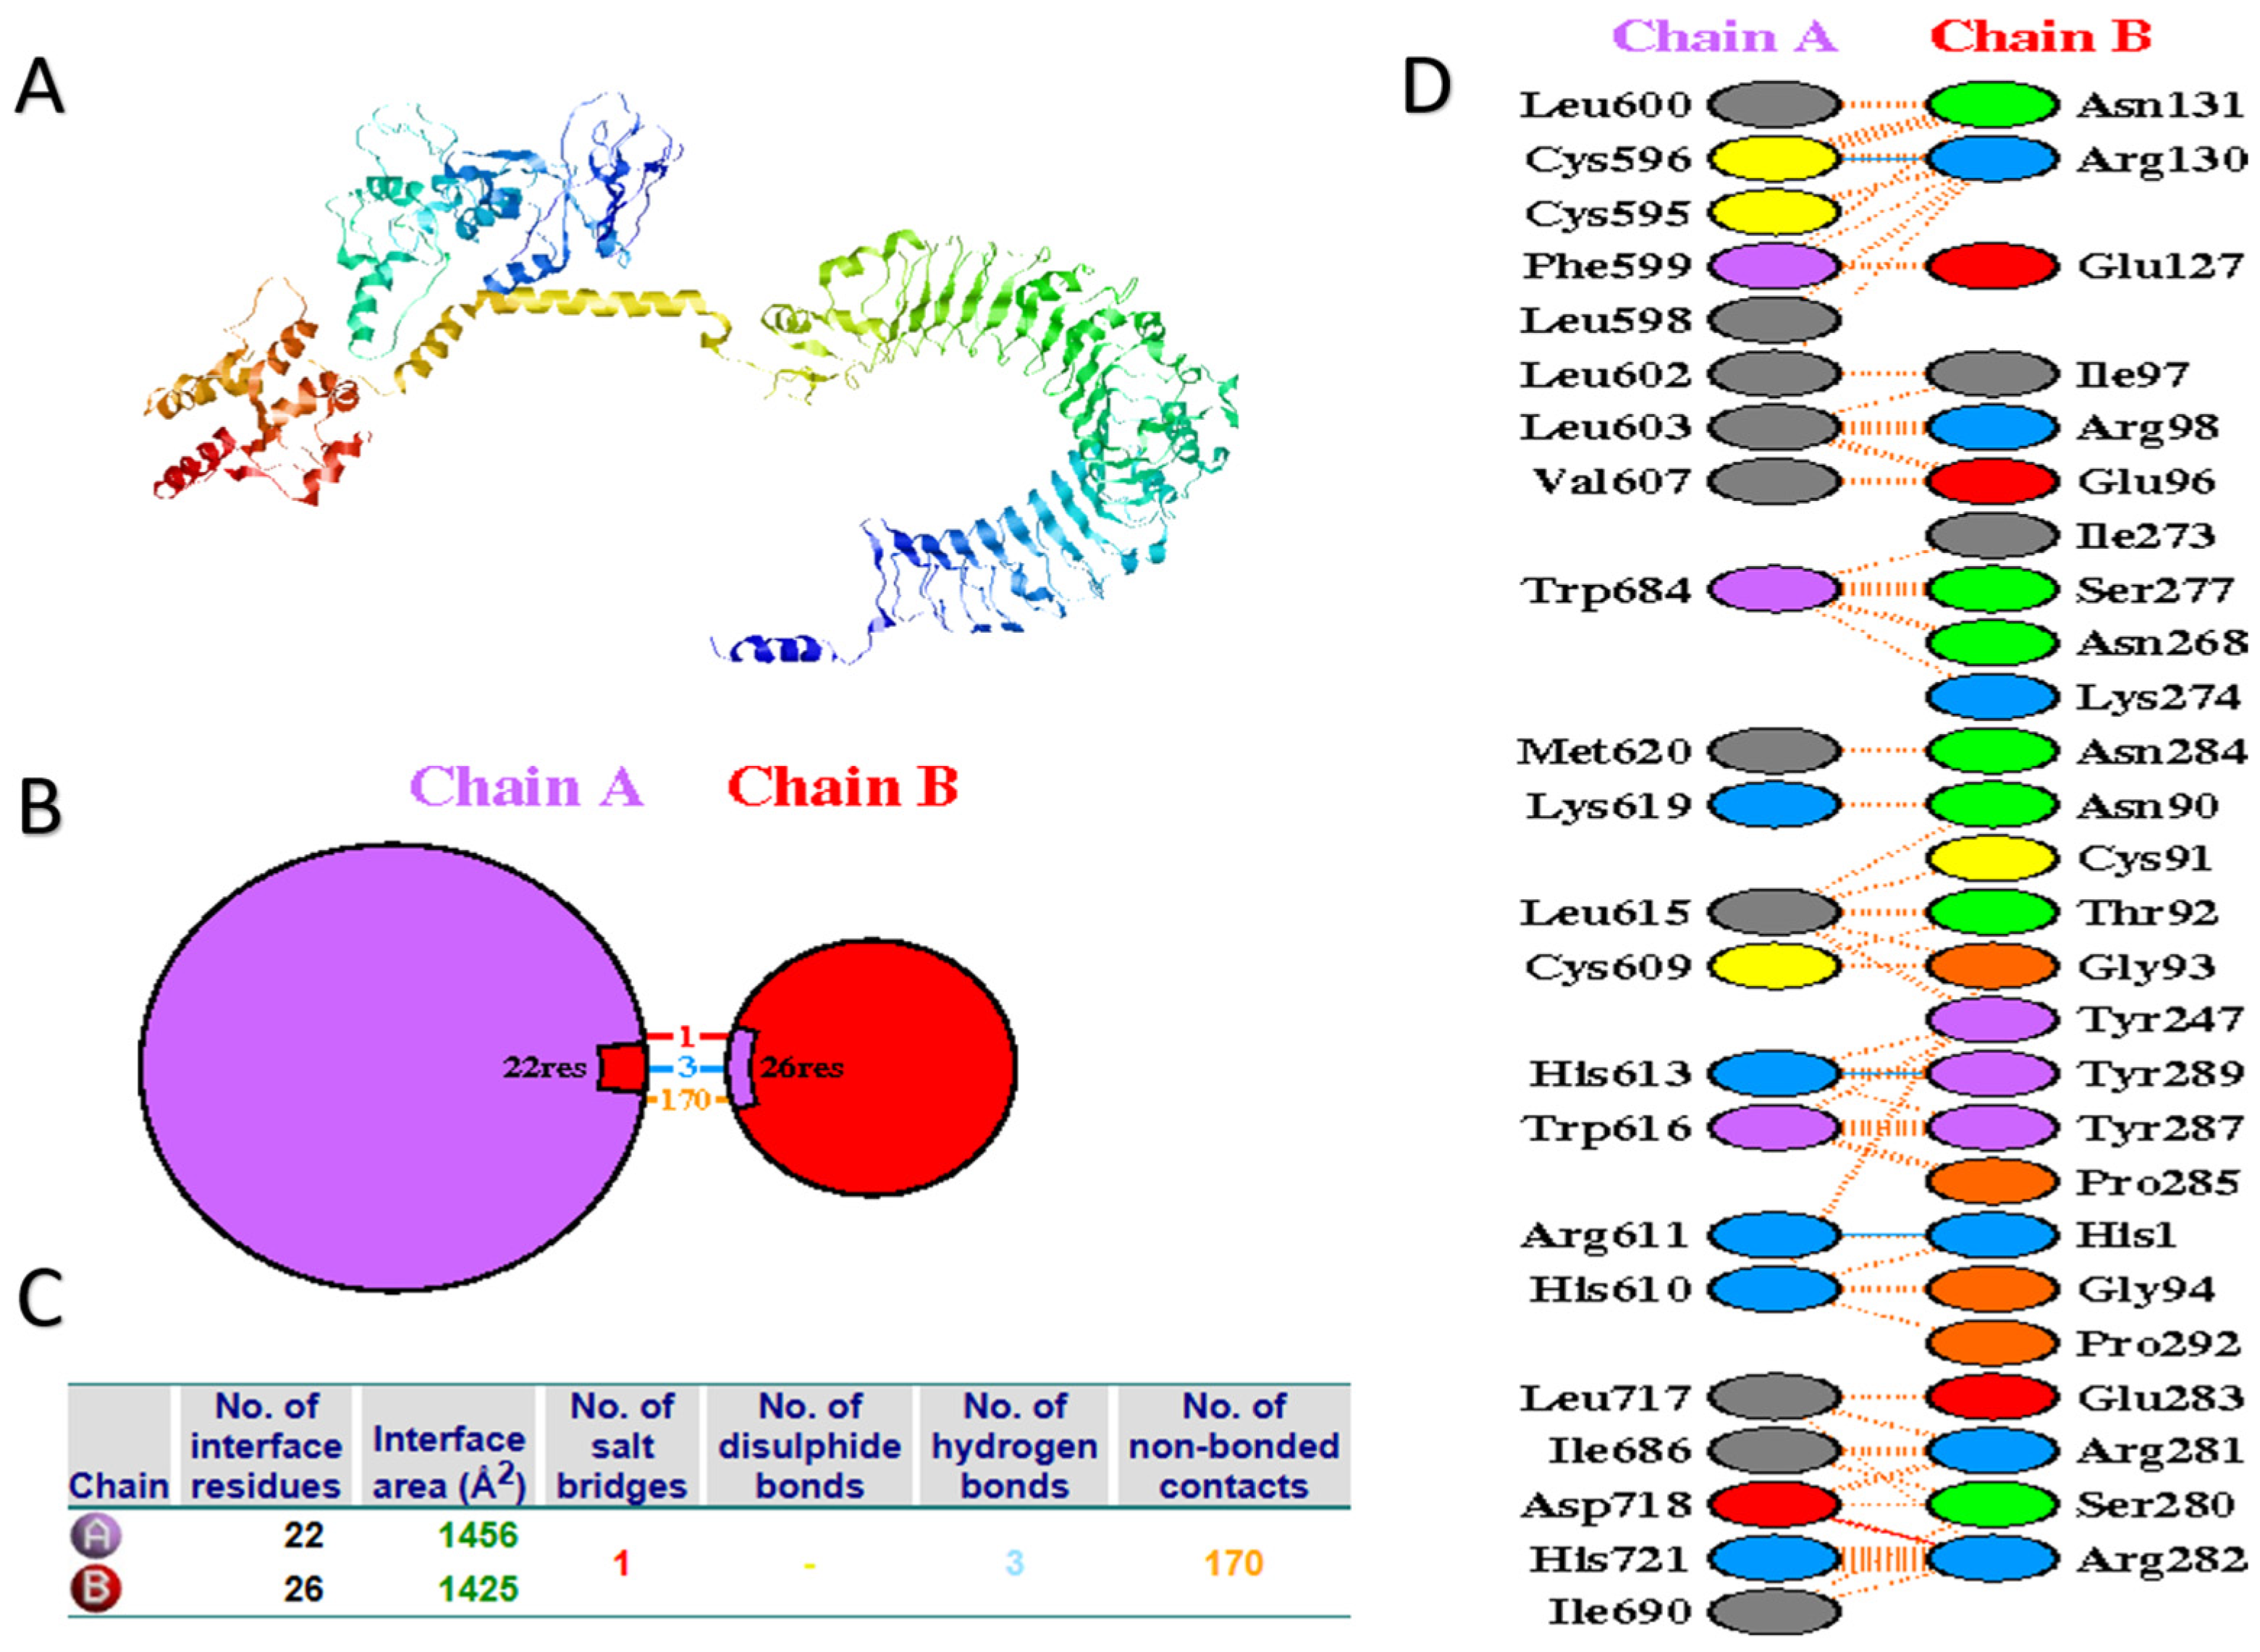
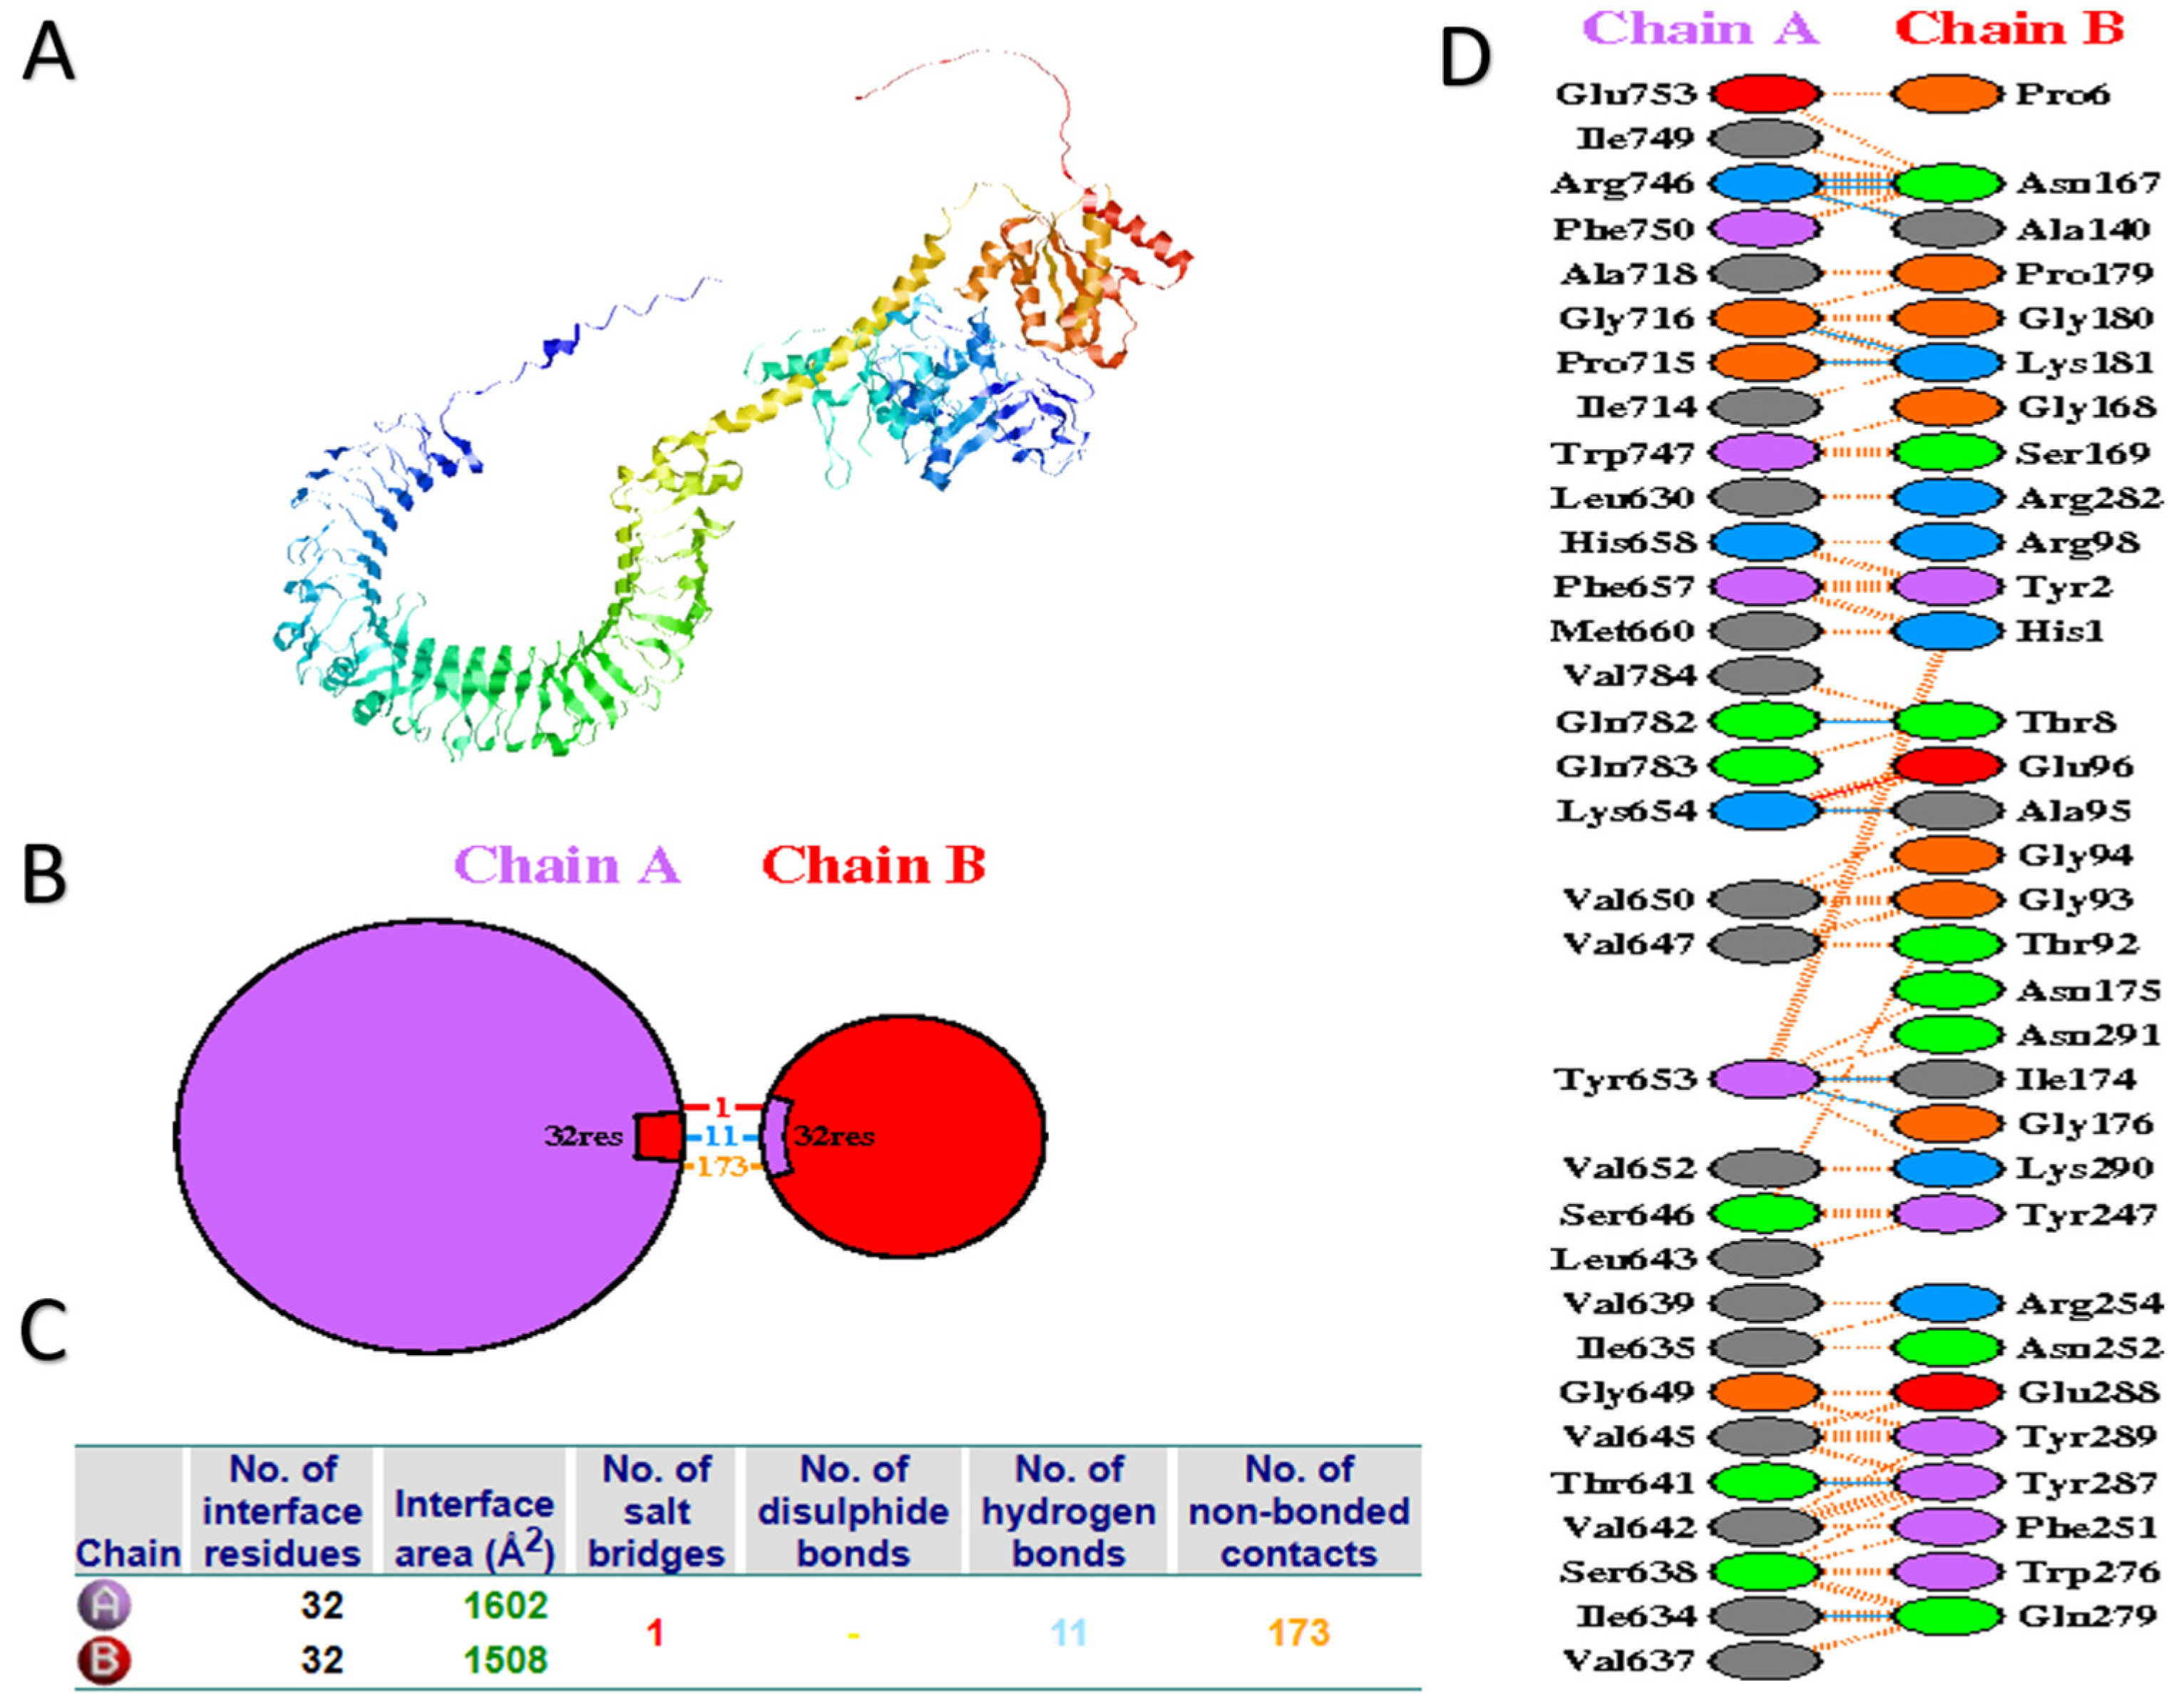
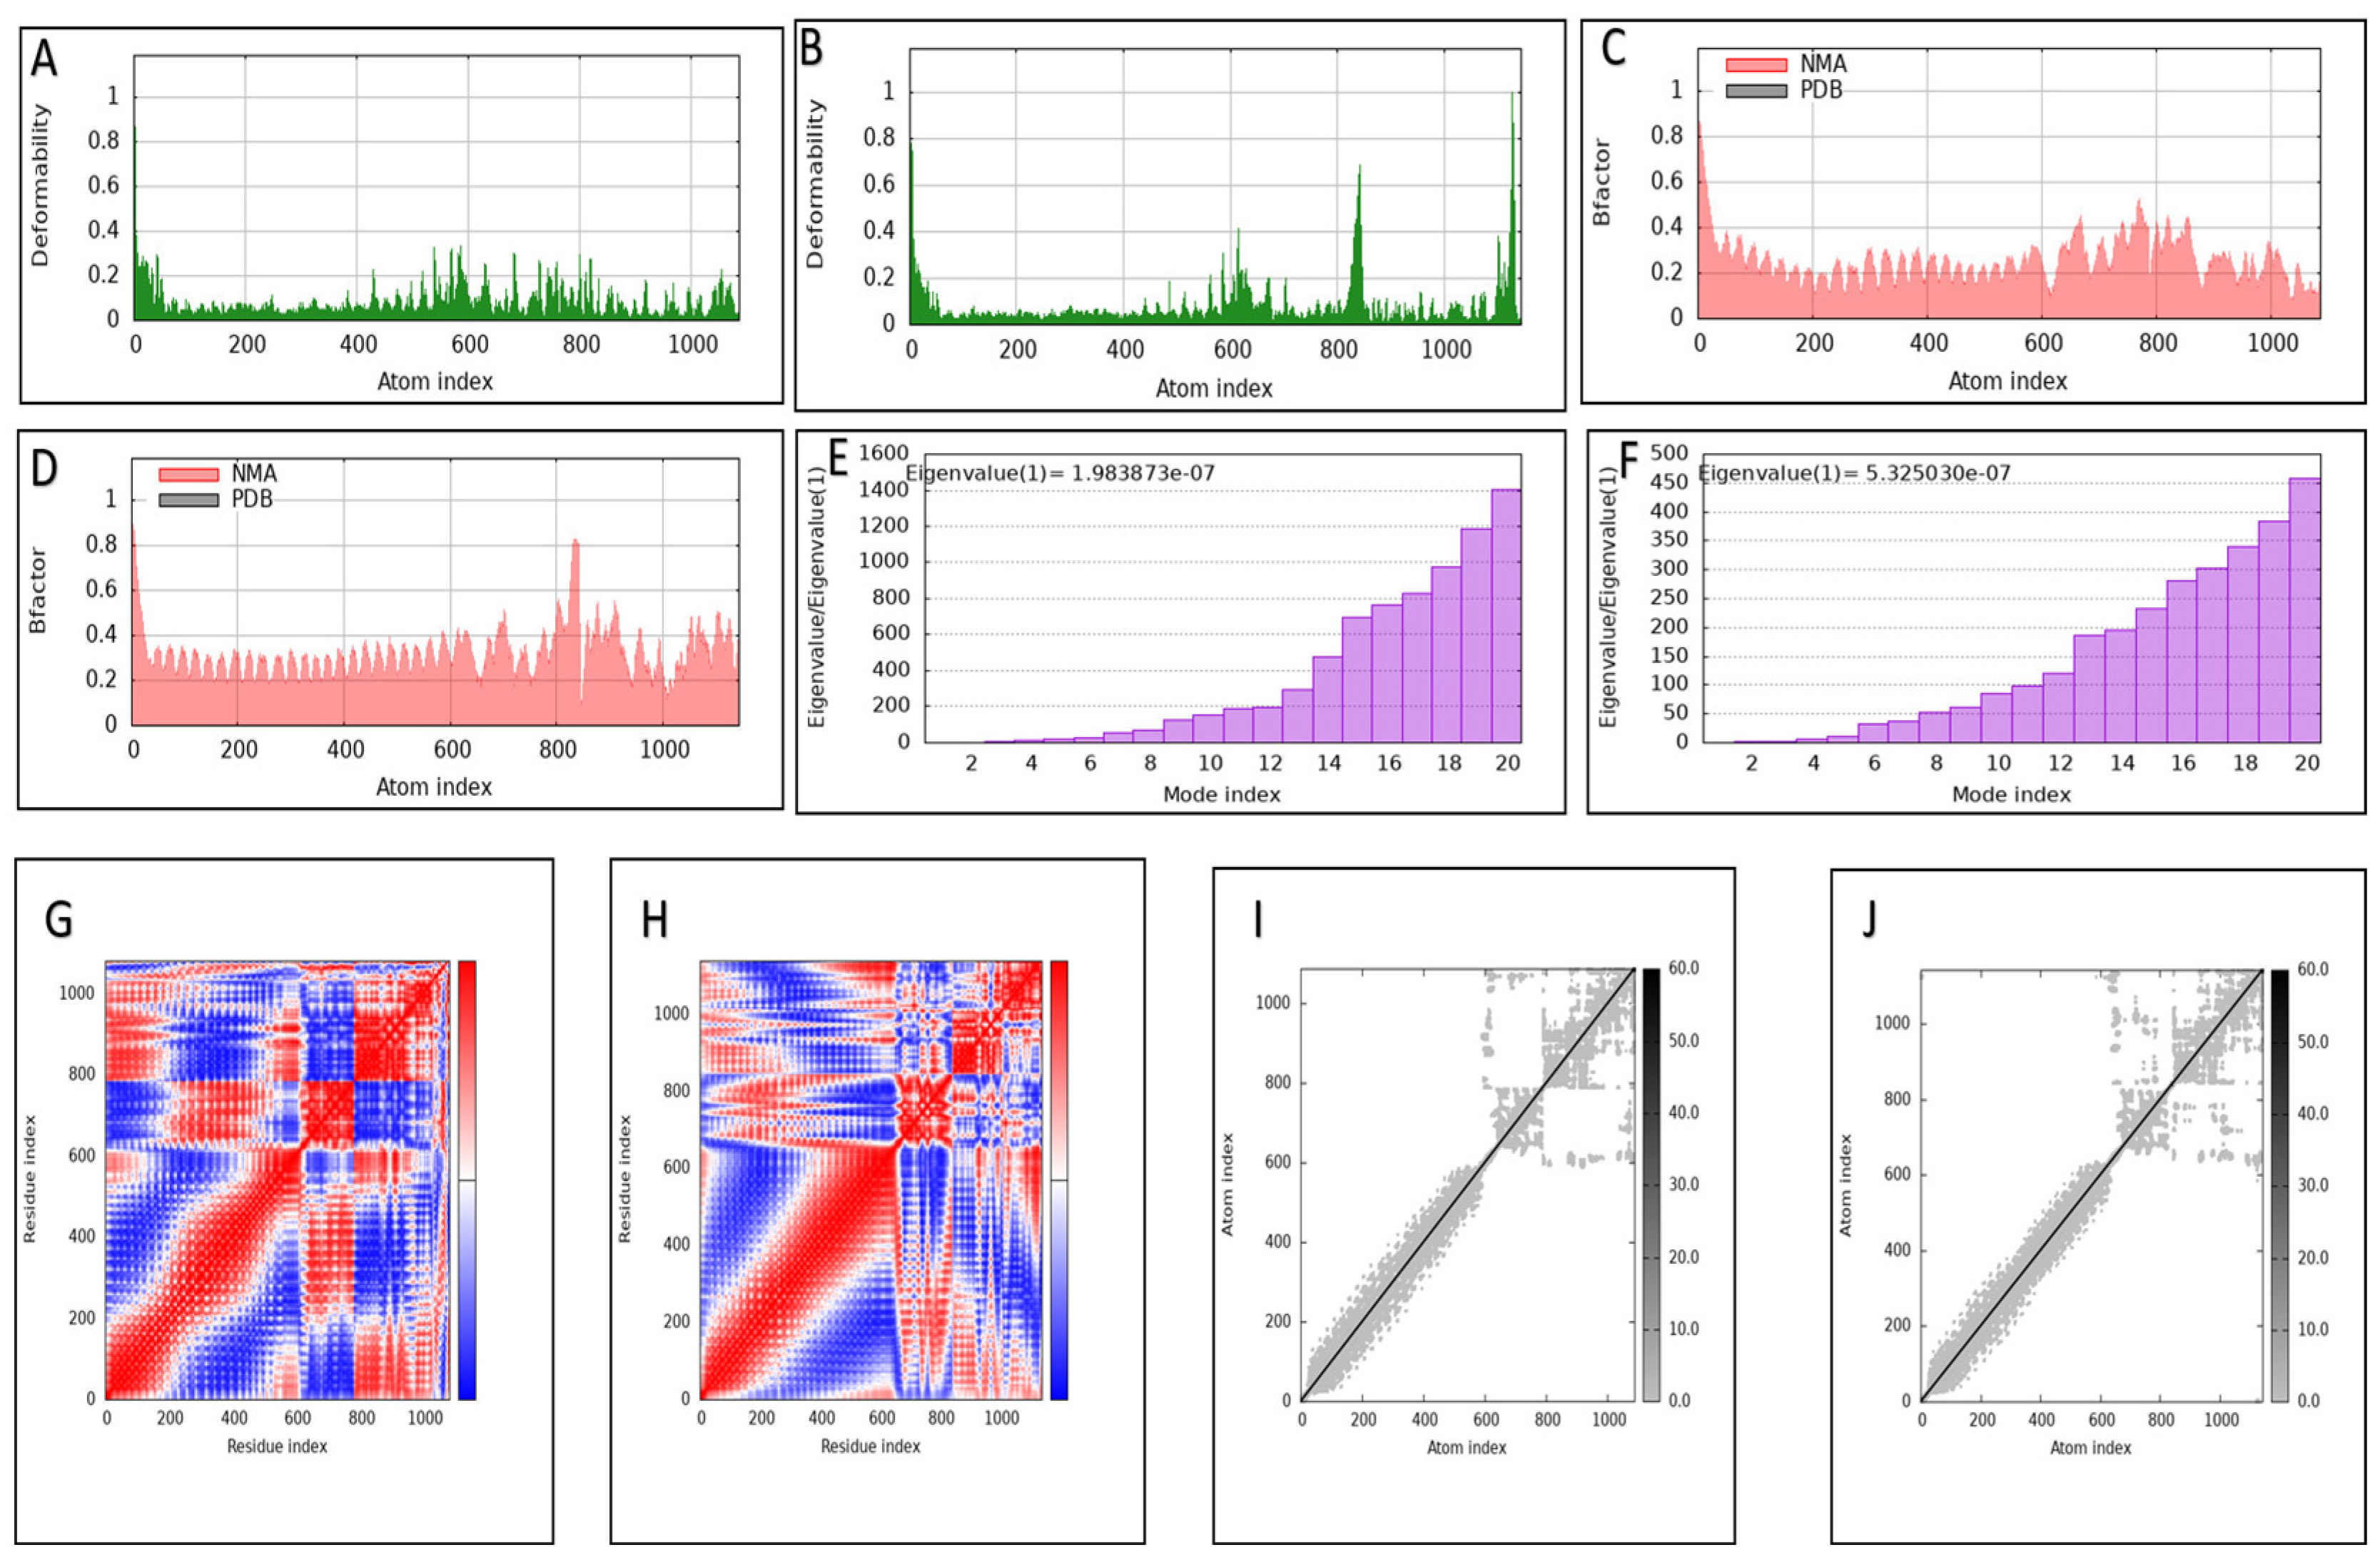
AS mentioned in method section the ClusPro v2.0 web server for molecular docking of the vaccine with the selected immune receptors (Bovine TLR2 and Bovine TLR4). The server predicted 10 possible complexes for each receptor. In the case of TLR2, model 1 was selected with the lowest energy score of −1127 (Figure 5A). While the lowest energy of model 1 of Vaccine–Bovine TLR4 was −1942.4 (Figure 6A). The PDBsum web server (https://www.ebi.ac.uk/thornton-srv/databases/pdbsum, accessed on: 12 February 2025) was used to predict the interacting residue within the complexes. As shown in Figure 5B–D, there are 22 and 26 residues in the interface, generating an interface area of 1456 and 1425 between the TLR2 and vaccine, respectively. The results show that the TLR2-Vaccine complex formed 1 salt bridge, 3 hydrogen bonds, and 170 nonbonding contacts. In the case of vaccine-TLR4, there are 32 and 32 residues in the interface, generating an interface area of 1605 and 1508 between the TLR4 and vaccine, respectively. One salt bridge, 11 hydrogen bonds, and 173 nonbonding contacts were found within the vaccine–TLR4 complex (Figure 6B–D). Normal mode analysis (NMA) of the vaccine–TLR complexes were predicted using the iMODS server. Figure 7A,B illustrated the deformability plot, which shows low deformity in the vaccine–TRL2 complex (Figure 7A) and vaccine–TLR4 complex (Figure 7B). This result predicted high stability and flexibility of the two complexes. A comparison between NMA and PDB for the vaccine–TLR was presented in Figure 7C,D. In both complexes, the similarity between predicted NMA and PDB refers to overall structural stability with localized flexible regions of the candidate construct. In Figure 7E,F, pure purple refers to individual variance with a clear exponential-like increase as the mode index increases. Both constructs are highly stable; however, vaccine—TLR4 has a more rigid and stable scaffold (Figure 7E) than vaccine-TLR2 (Figure 7F).
Figure 5.
Molecular interaction between bovine TLR2 and the vaccine candidate. (A): 3D structural diagram of molecular docking of the proposed vaccine with TLR2 receptor. (B): Schematic diagram for TRL2 and the vaccine candidate interaction. (C): Interface statistics. (D): Overview interaction between TLR2 and the vaccine showing residues and bonds.
Figure 6.
Molecular interaction between bovine TLR4 and the vaccine candidate. (A): 3D structural diagram of molecular docking of the proposed vaccine with TLR2 receptor. (B): Schematic diagram for TLR4 and the vaccine candidate interaction. (C): Interface statistics. (D): Overview interaction between TLR4 and the vaccine showing residues and bonds.
Figure 7.
NMA modeling of Vaccine–Bovine TLR complexes. (A,B): Modeling the deformability of the main chain for TLR2 and TLR4. (C): B-factor values (Vaccine–TLR2). (D): B-factor values (Vaccine–TLR4). (E,F): Eigenvalues value the complex (Vaccine with TLR2 and TLR4, respectively). (G,H): Graph representing covariance of the complex (Vaccine–TLR2 and TLR4). (I,J): Model of elastic network (TLR2 and TLR4, respectively).
Covariance matrix was represented in Figure 7G,H where the correction between residue pairs has been detected. Finally, the combination of the vaccine within the receptor was predicted using the elastic network model (Figure 7I,J). Overall, the results predicted that the tested complexes are strong and stable.
3.8. Immune Simulation of the Proposed Vaccine
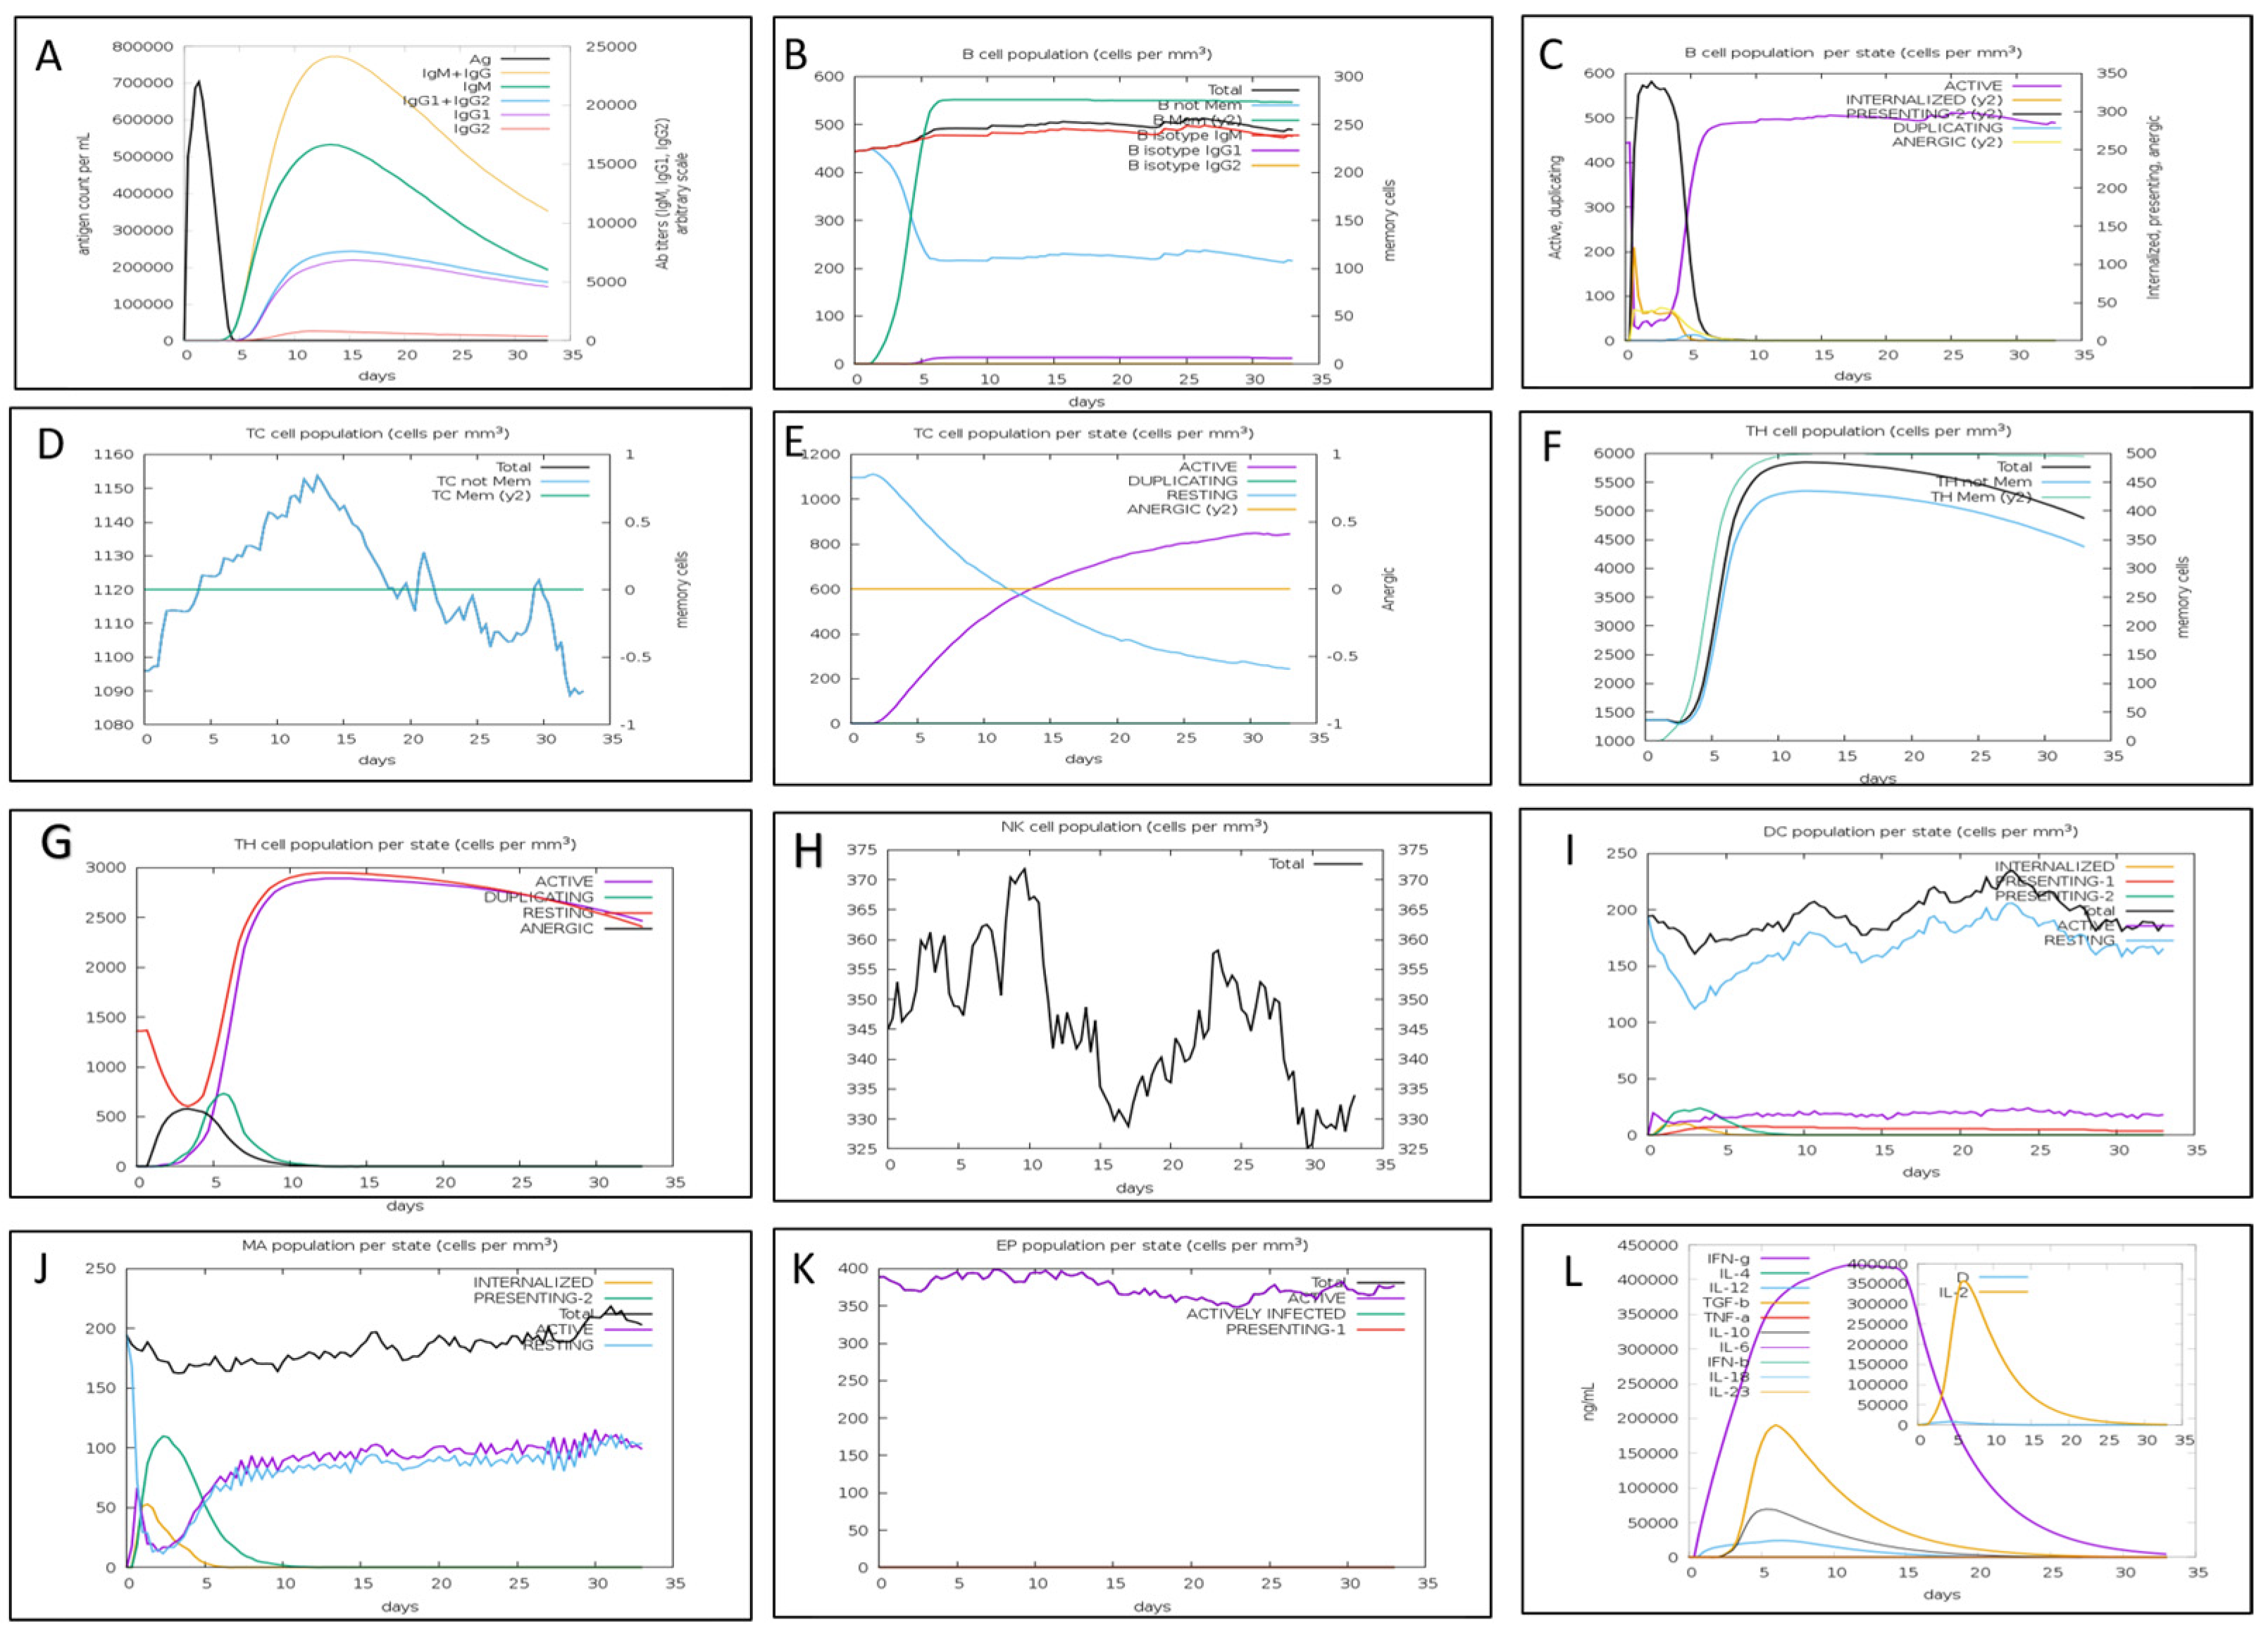
The C-ImmSim web server was used to simulate immune responses after three injections of the proposed vaccine. The results in general show that the vaccine computationally is able to induce humoral and cellular responses (Figure 8). Figure 8A shows a rapid and significant increase in the titer of the antibody after injection of the proposed vaccine. The number of Memory B cells was also increased after immunization, reaching 700 cells/mm3 and remaining high, which suggests long-term potential protection of the proposed vaccine (Figure 8B). The concentration of activated B cells was 480–510 cells/mm3 (Figure 8C). In the same manner, TH and TC were also notably increased in response to the injection of the proposed vaccine where the helper T cell enhanced immune regulation and coordination, while performing T cytotoxic targeting and elimination of the infected cells (Figure 8D–G).
Figure 8.
Chimeric multi-epitope vaccine immune-simulation. (A): Level of immunoglobulin after immunization. (B): B-cell population. (C): B-cell population per state. (D): T cytotoxic cell population. (E): T cytotoxic cell per state. (F): T helper cell population. (G): T helper cell population per state. (H): Nature killer population. (I): Dendritic cell per state. (J): Macrophage population per state. (K): Epithelial cell population per state. (L): Cytokines and interleukins concentration.
It should be noted that the C-ImmSim server is not specifically parameterized for bovine immunology (e.g., BoLA restriction) nor for scenarios involving pregnant dams, maternal antibody transfer, or neonatal protection via colostrum. Therefore, these simulation results should be interpreted as qualitative predictions of general immune trends. Experimental validation in the target cattle population is required to confirm actual immunogenicity and protective efficacy.
3.9. mRNA Vaccine Structure and In Silico Codon Optimization
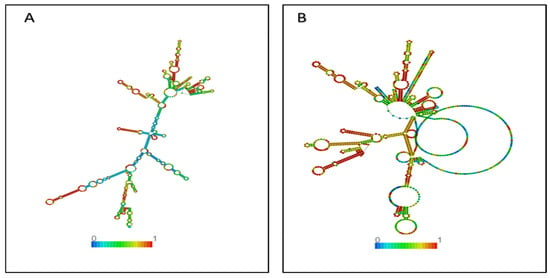
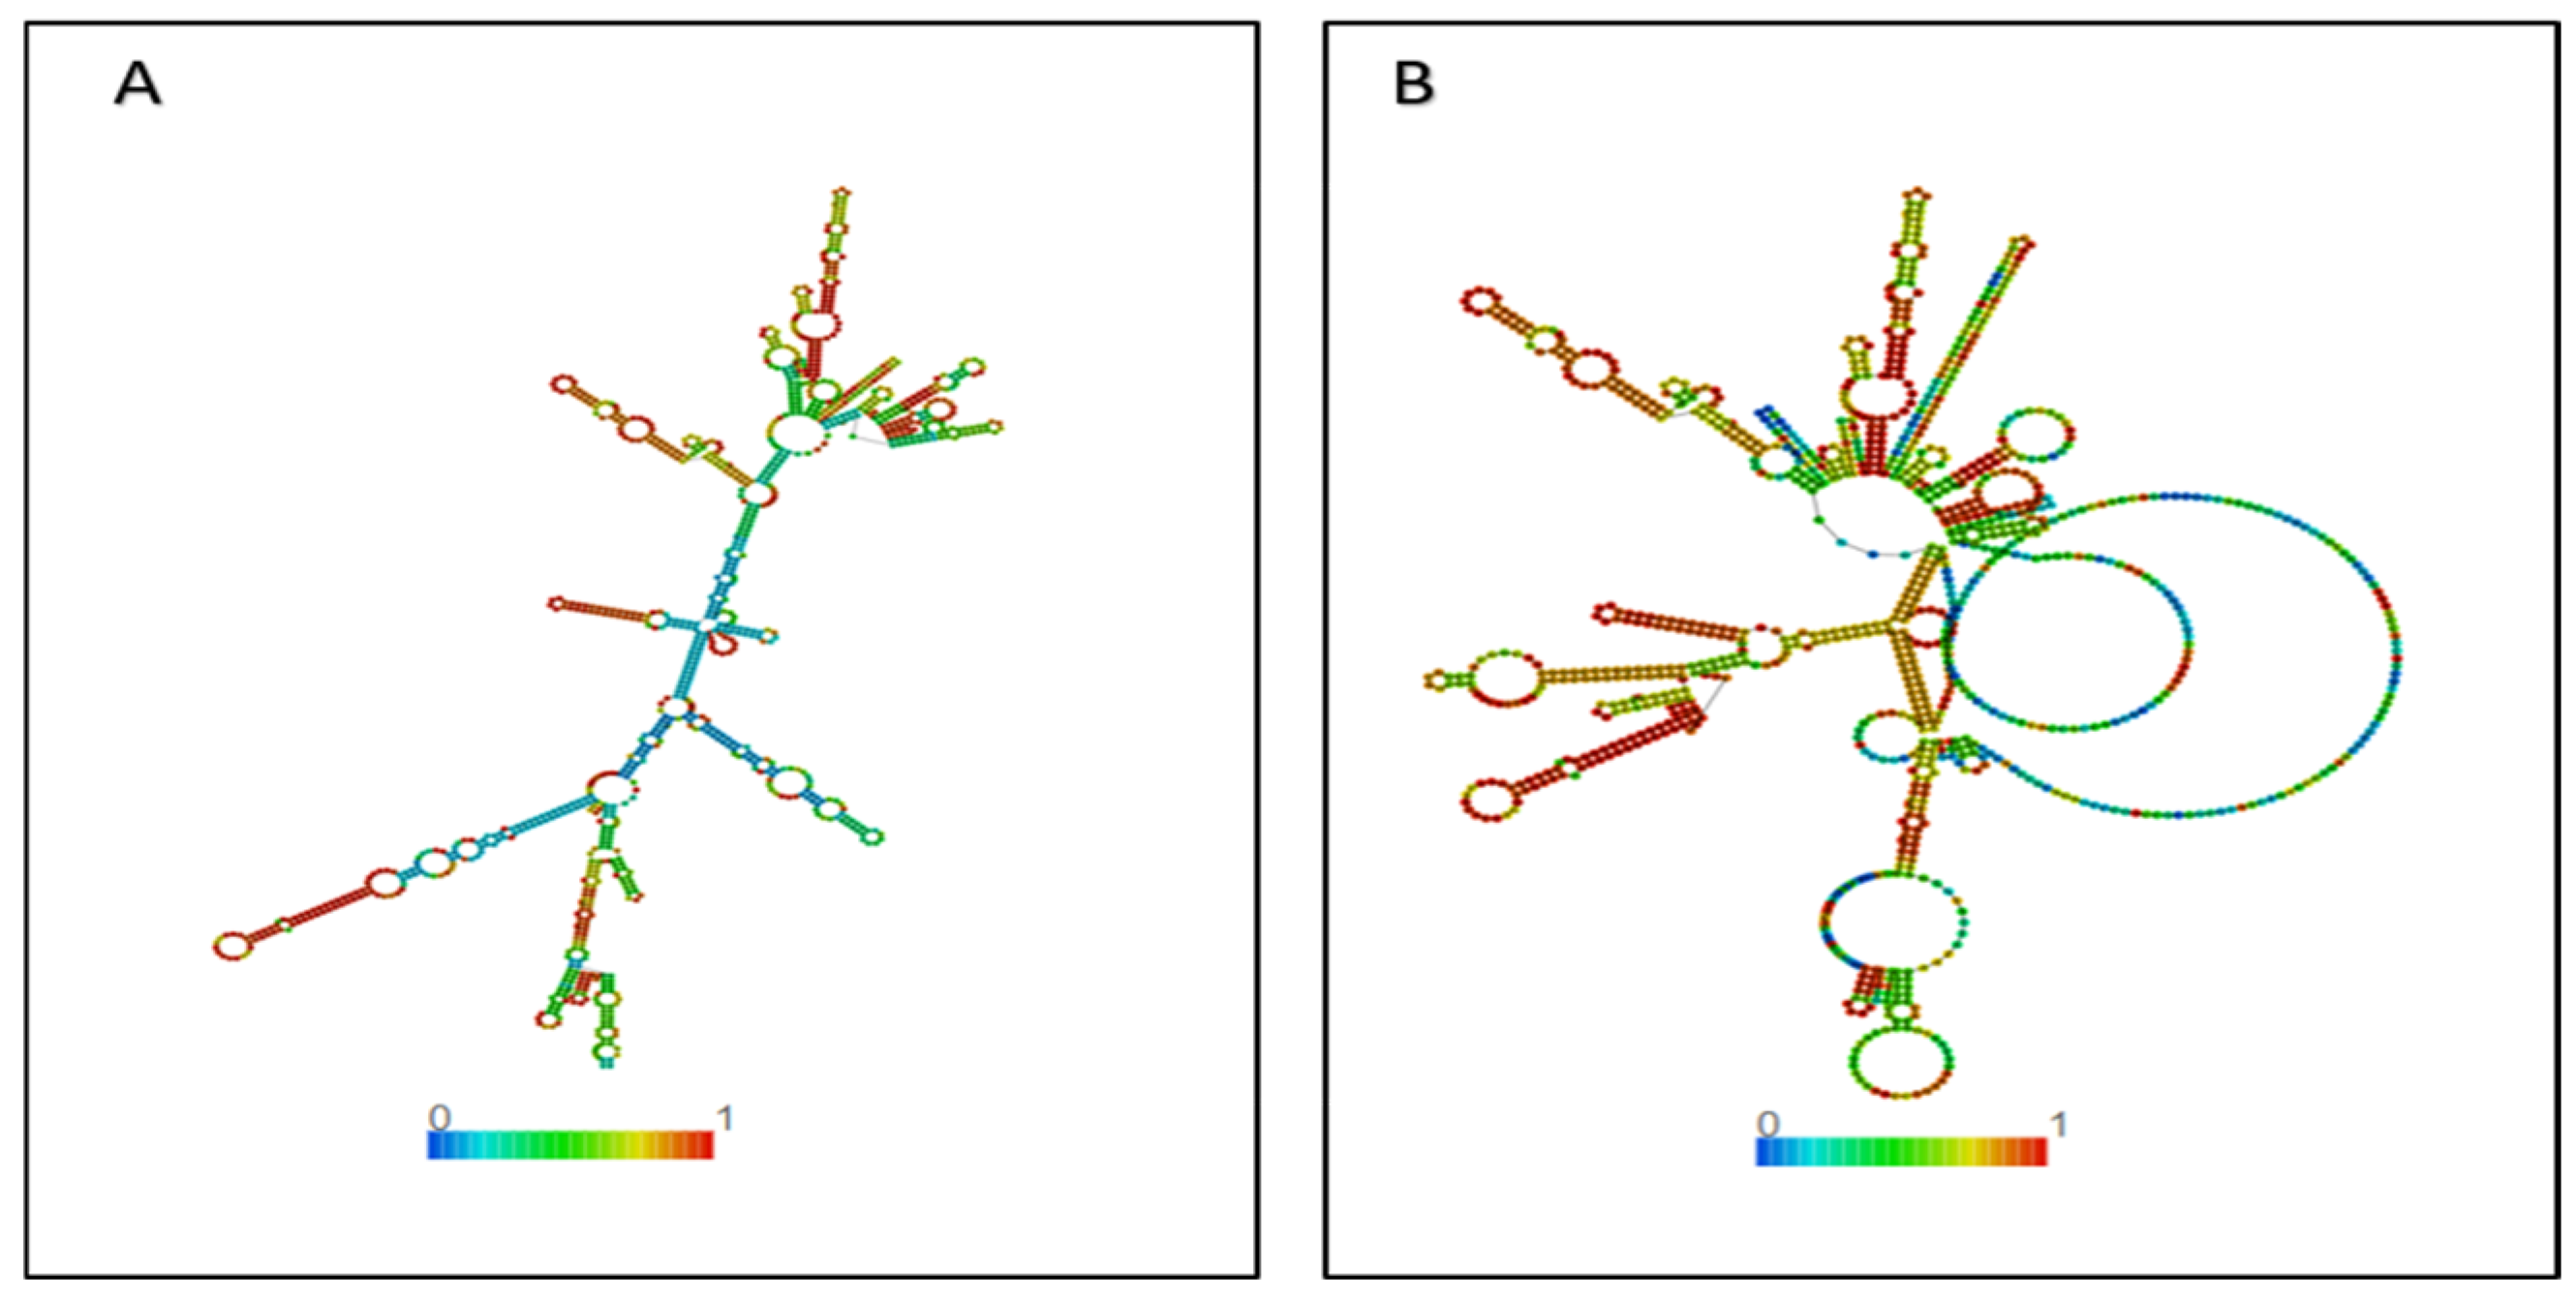
Codon optimization for the sequence of the proposed vaccine was performed using the VectorBuilder 3.5 (USA) web server. Optimization of the sequence was performed using default setting, except the target host was changed to cow. GC content of the sequence became 59.68% after optimization, while CAI was 0.87, which overall results in a high translation possibility of RNA within the cow body. The RNAfold web server was used to predict the structural arrangement of the RNA molecule. The results show that the centroid secondary structure free energy was −240.10 kcal/mol (Figure 9B), while the thermodynamic ensemble was −321.42 kcal/mol (Figure 9A). In general, the result of thermodynamic robustness predicts that tested modeled RNA was good and stable.
Figure 9.
Structure of the modeled RNA molecule. (A): Ideal secondary configuration. (B): Central secondary arrangement of the RNA.
4. Discussion
Neonatal calf diarrhea is a multifactorial disease mainly caused by Bovine Coronavirus (BCoV), Bovine Rotavirus (BRV) and Escherichia coli K99, leading to severe economic losses and high mortality during the first weeks of life [2]. Infected calves with more than one NCD cause are frequently reported in many recent studies. Furthermore, these studies suggest that clinical signs of mixed infection are more severe than single types of infection with NCD causes [23]. Gabriele and his colleagues summarized the research that focuses on the development of a vaccine against the main NCD pathogens; they found that most proposed vaccines targeted E. coli, Bovine Rotavirus, Salmonella, Bovine Coronavirus and Bovine Viral Diarrhea Virus. However, multi-antigen type vaccines that target theses pathogens are not yet valid [24]. The lack of validity for commercial vaccines that target the main causes of NCD make control of the disease is difficult.
In recent years, several multi-epitope-based vaccines were designed against wide ranges of infectious disease agents using an immunoinformatic tool. Promising results were gotten from evaluating some of these vaccines in terms of their ability to induce immune response within animal models or provide protective immunity against the target agent. However, multifactorial disease involving more than one pathogen are urgently needed for an innovative vaccine that provides protection across strains [25].
Recent advances in immunoinformatic tools along with structural biology leads to the possible use of a fusion strategy to develop a multivalent vaccine against heterogeneous pathogens. Yang and his colleagues utilized this strategy to design a chimeric multi-epitope vaccine targeting influenza A and Mycoplasma pneumonia [26], while Mohammadipour and his colleagues targeted SARS-CoV-2 and influenza viruses using a single multi-epitope vaccine [27]. During the COVID-19 outbreak, numerous studies focused on designing and evaluating mRNA vaccines against several important pathogens. mRNA vaccines offer multiple advantages, including rapid and straightforward development, relatively low production costs, and a strong safety profile, as they do not pose a risk of genomic integration [28].
In this study, we, for the first time, engineered a single vaccine based on mRNA targeting BCV, BRV and E. coli K99. Immunoinformatic and molecular modeling approaches were used to design an immunoinformatic evaluation of the proposed vaccine. According to their role in pathogenicity and initiation of infection, three proteins of targeted pathogens were selected to construct the vaccine candidate. The proposed vaccine consists of 15 epitopes in total, with the addition of building an adjuvant to increase the immune response. Furthermore, the proposed vaccine was tested using set of immunoinformatic tools to be able to predict its antigenicity, stability, and its ability to induce an immune response within the target host. Structural modelling confirms that the vaccine is conformationally stable and is highly likely to be recognized by immune cells. Furthermore, docking analysis show that the proposed vaccine is able to interact with Bovine TRL2 and Bovine TLR4 at low energy. Moreover, immune simulation predicts the vaccine’s ability to induce a strong humoral and cellular response.
VP6 was selected over classical neutralization targets (VP4/VP7) because it is the most conserved and immunogenic rotavirus structural protein, capable of inducing cross-protective immunity across different strains via intracellular neutralization—a mechanism where anti-VP6 antibodies internalized into infected cells bind to the middle capsid during virus uncoating, thereby inhibiting viral replication. This strategy ensures broad efficacy against diverse BRV strains in the context of a multi-pathogen vaccine [29,30].
The 5′ UTR from human β-globin (HBB) and the 3′ UTR from bovine growth hormone (bGH) were selected to enhance the translation and mRNA stability. A 120-nucleotide poly(A) tail, a Cap1 structure (e.g., CleanCap® AG, Tinzyme, China), and full substitution of uridine with N1-methylpseudouridine (m1Ψ) were incorporated to maximize translation and reduce innate immune recognition. For in vivo delivery, we propose formulating the mRNA in lipid nanoparticles (LNPs) composed of SM-102/DSPC/cholesterol/PEG2000-DMG (50:10:38.5:1.5 molar ratio), which would produce particles of 80–100 nm. The rationale for administering this mRNA-LNP vaccine to pregnant cattle is that vaccination of dams induces high-titer IgG antibodies transferred via colostrum, providing immediate protection to calves during the first 10 days of life—the period of highest NCD susceptibility. Direct neonatal vaccination is hindered by immune immaturity and the rapid onset of diarrhea. mRNA platforms are particularly suitable for pregnant cattle because they carry no risk of reversion to virulence, do not require potentially abortifacient adjuvants, and are rapidly degraded without genomic integration, as demonstrated by their safe use in human pregnancy [31].
Overall, the proposed vaccine can be a promising option for the control of NCD. A limitation of the present study is the use of representative BoLA alleles derived from the Holstein breed rather than breed-specific frequency data; thus, the epitope set should be validated experimentally in the target cattle population before field application.
Also, the RpfE adjuvant has two limitations: (1) potential cross-reaction with bovine tuberculosis diagnostic tests (e.g., IFN-γ release assays, skin tests) due to its mycobacterial origin, which future studies should address using DIVA strategies; and (2) the lack of a control construct without RpfE in this in silico study, so comparative docking/immune simulations are needed to confirm that observed TLR4 binding and immunogenicity are driven by the NCD epitopes rather than the adjuvant alone. Future experimental work should include such a control.
Several wet laboratory studies are required to confirm several issues including expression of the mRNA-based vaccine within the host, safety of the vaccine, ability to induce specific immunity against the selected pathogen and protective efficiency against natural experimental infection with the NCD pathogen.
5. Conclusions
In this study, immunoinformatic and molecular modeling approaches were used to design a chimeric multi-epitope mRNA vaccine targeting BCoV, BRV, and E. coli K99. In silico analyses suggest the vaccine is stable and may induce immune responses, but limitations include the use of representative BoLA alleles, a non-bovine simulation platform, and the absence of a control construct without the RpfE adjuvant. Therefore, conclusions regarding protective immunity are preliminary, and laboratory experiments are needed to confirm efficacy and safety.
Supplementary Materials
The following supporting information can be downloaded at: https://www.mdpi.com/article/10.3390/immuno6030048/s1.
Author Contributions
A.A. and B.S.K. Identified highly antigenic proteins within the BCV, BRV and E coli K99. M.H. and D.A.: Software and analysis. All authors have read and agreed to the published version of the manuscript.
Funding
This research received no external funding.
Institutional Review Board Statement
Not applicable.
Informed Consent Statement
Not applicable.
Data Availability Statement
Data available on request from corresponding author Amjed Alsultan.
Acknowledgments
The authors thank the Deanship of the Veterinary Medicine College, Al-Qadisiyah University for providing the necessary facilities for this study.
Conflicts of Interest
The authors declare that there are no conflicts of interest.
References
- Foster, D.M.; Smith, G.W. Pathophysiology of diarrhea in calves. Vet. Clin. N. AM. Food Anim. Pract. 2009, 25, 13–36. [Google Scholar] [CrossRef] [Scilit]
- Cho, Y.I.; Yoon, K.J. An overview of calf diarrhea—Infectious etiology, diagnosis, and intervention. J. Vet. Sci. 2014, 15, 1–17. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Kostanić, V.; Kunić, V.; Prišlin Šimac, M.; Lolić, M.; Sukalić, T.; Brnić, D. Comparative insights into acute gastroenteritis in cattle caused by bovine rotavirus A and bovine coronavirus. Vet. Sci. 2024, 11, 671. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Maier, G.U.; Breitenbuecher, J.; Gomez, J.P.; Samah, F.; Fausak, E.; Van Noord, M. Vaccination for the prevention of neonatal calf diarrhea in cow-calf operations: A scoping review. Vet. Anim. Sci. 2022, 15, 100238. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Pardi, N.; Hogan, M.J.; Porter, F.W.; Weissman, D. mRNA vaccines—A new era in vaccinology. Nat. Rev. Drug Discov. 2018, 17, 261–279. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Pumchan, A.; Proespraiwong, P.; Sawatdichaikul, O.; Phurahong, T.; Hirono, I.; Unajak, S. Computational design of novel chimeric multiepitope vaccine against bacterial and viral disease in tilapia (Oreochromis sp.). Sci. Rep. 2024, 14, 14048. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Leong, K.Y.; Tham, S.K.; Poh, C.L. Revolutionizing immunization: A comprehensive review of mRNA vaccine technology and applications. Virol. J. 2025, 22, 71. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Zhang, H.; Yuan, X.; He, Y.; Chen, Y.; Hu, C.; Chen, J.; Zhang, L.; Chen, X.; Guo, A. A novel multiepitope fusion antigen as a vaccine candidate for the prevention of enterotoxigenic E. Coli-Induc. Calf Diarrhea. Vaccines 2024, 12, 457. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Zhang, Q.; Wang, P.; Kim, Y.; Haste-Andersen, P.; Beaver, J.; Bourne, P.E.; Bui, H.H.; Buus, S.; Frankild, S.; Greenbaum, J.; et al. Immune epitope database analysis resource (IEDB-AR). Nucleic Acids Res. 2008, 36, W513–W518. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Vita, R.; Mahajan, S.; Overton, J.A.; Dhanda, S.K.; Martini, S.; Cantrell, J.R.; Wheeler, D.K.; Sette, A.; Peters, B. The immune epitope database (IEDB): 2018 update. Nucleic Acids Res. 2019, 47, D339–D343. [Google Scholar] [PubMed]
- Nassiry, M.R.; Eftekhar Shahroodi, F.; Mosafer, J.; Mohammadi, A.; Manshad, E.; Ghazanfari, S.; Mohammad Abadi, M.R.; Sulimova, G.E. Analysis and frequency of bovine lymphocyte antigen (BoLA-DRB3) alleles in Iranian Holstein cattle. Russ. J. Genet. 2005, 41, 664–668. [Google Scholar] [CrossRef] [Scilit]
- Bui, H.H.; Sidney, J.; Dinh, K.; Southwood, S.; Newman, M.J.; Sette, A. Predicting population coverage of T-cell epitope-based diagnostics and vaccines. BMC Bioinform. 2006, 7, 153. [Google Scholar] [CrossRef] [Scilit]
- Doytchinova, I.A.; Flower, D.R. VaxiJen: A server for prediction of protective antigens, tumour antigens and subunit vaccines. BMC Bioinform. 2007, 8, 4. [Google Scholar] [CrossRef] [Scilit]
- Dimitrov, I.; Bangov, I.; Flower, D.R.; Doytchinova, I. AllerTOP v. 2—A server for in silico prediction of allergens. J. Mol. Model. 2014, 20, 2278. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Chen, X.; Zaro, J.L.; Shen, W.C. Fusion protein linkers: Property, design and functionality. Adv. Drug Deliv. Rev. 2013, 65, 1357–1369. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Gasteiger, E.; Hoogland, C.; Gattiker, A.; Duvaud, S.E.; Wilkins, M.R.; Appel, R.D.; Bairoch, A. Protein identification and analysis tools on the ExPASy server. In The Proteomics Protocols Handbook 22 March 2005; Humana Press: Totowa, NJ, USA, 2005; pp. 571–607. [Google Scholar]
- McGuffin, L.J.; Bryson, K.; Jones, D.T. The PSIPRED protein structure prediction server. Bioinformatics 2000, 16, 404–405. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Roy, A.; Kucukural, A.; Zhang, Y. I-TASSER: A unified platform for automated protein structure and function prediction. Nat. Protoc. 2010, 5, 725–738. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Heo, L.; Park, H.; Seok, C. GalaxyRefine: Protein structure refinement driven by side-chain repacking. Nucleic Acids Res. 2013, 41, W384–W388. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Lovell, S.C.; Davis, I.W.; Arendall, W.B., 3rd; De Bakker, P.I.; Word, J.M.; Prisant, M.G.; Richardson, J.S.; Richardson, D.C. Structure validation by Cα geometry: ϕ, ψ and Cβ deviation. Proteins Struct. Funct. Bioinform. 2003, 50, 437–450. [Google Scholar] [CrossRef] [Scilit]
- Guo, W.; Wang, X.; Hu, J.; Zhang, B.; Zhao, L.; Zhang, G.; Qi, J.; Wei, Z.; Bao, Y.; Tian, M.; et al. In silico design of a multi-epitope vaccine against Mycobacterium avium subspecies paratuberculosis. Front. Immunol. 2025, 16, 1505313. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Rapin, N.; Lund, O.; Bernaschi, M.; Castiglione, F. Computational immunology meets bioinformatics: The use of prediction tools for molecular binding in the simulation of the immune system. PLoS ONE 2010, 5, e9862. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Brunauer, M.; Roch, F.F.; Conrady, B. Prevalence of worldwide neonatal calf diarrhoea caused by bovine rotavirus in combination with bovine coronavirus, Escherichia coli K99 and Cryptosporidium spp.: A meta-analysis. Animals 2021, 11, 1014. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Alawadi, Z.I.; Alsultan, A.; Alsallami, D.; Alnomasy, S.F.; Alqasmi, M.; Almufarriji, F.M.; Alotaibi, B.S.; Mazhari, B.B.; Alenazy, R. Designing a multiepitope mRNA-based vaccine against enterotoxigenic Escherichia coli infection in calves: Immunoinformatics and molecular modeling approach. Open Vet. J. 2024, 14, 1417. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Yang, Z.; Li, Y.; Chen, L.; Jia, S. Designing a broad-spectrum multi-epitope vaccine against influenza A and Mycoplasma pneumoniae: An immunoinformatics approach. Front. Public Health 2026, 14, 1671035. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Mohammadipour, S.; Tavakkoli, H.; Fatemi, S.N.; Sharifi, A.; Mahmoudi, P. Designing a multi-epitope universal vaccine for concurrent infections of SARS-CoV-2 and influenza viruses using an immunoinformatics approach. BMC Infect. Dis. 2025, 25, 688. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Kim, S.C.; Sekhon, S.S.; Shin, W.R.; Ahn, G.; Cho, B.K.; Ahn, J.Y.; Kim, Y.H. Modifications of mRNA vaccine structural elements for improving mRNA stability and translation efficiency. Mol. Cell. Toxicol. 2022, 18, 1–8. [Google Scholar] [PubMed]
- Krasnikova, S.I.; Sharabrin, S.V.; Kisakov, D.N.; Borgoyakova, M.B.; Starostina, E.V.; Tigeeva, E.V.; Makarova, K.P.; Yakovlev, V.A.; Ivanova, K.I.; Rudometov, A.P.; et al. Effect of Modified Nucleotides on the Immunogenicity of the H1N1 Influenza mRNA Vaccine. Bull. Exp. Biol. Med. 2025, 180, 603–608. [Google Scholar] [CrossRef] [Scilit]
- Wang, W.; Feng, S.; Ye, Z.; Gao, H.; Lin, J.; Ouyang, D. Prediction of lipid nanoparticles for mRNA vaccines by the machine learning algorithm. Acta Pharm. Sin. B 2022, 12, 2950–2962. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Hu, X.; Huang, J.; Cai, Y.; Zhang, W.; Cheng, Y. Bovine Viral Diarrhea Virus and Vaccine Protection Strategies. Vet. Sci. 2026, 13, 180. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Chaudhary, N.; Weissman, D.; Whitehead, K.A. mRNA vaccines for infectious diseases: Principles, delivery and clinical translation. Nat. Rev. Drug Discov. 2021, 20, 817–838. [Google Scholar] [CrossRef] [Scilit] [PubMed]

Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2026 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license.