


Abstract
Pladienolide B (1), a macrocyclic polyketide isolated from Streptomyces platensis Mer-11107, displays potent antiproliferative and antitumor activities through its interaction with the SF3b subunit of the spliceosome. Herein, we report a divergent, stereoselective synthetic route to access both the C1–C8 and C9–C13 fragments of the core moiety of Pladienolide B (1) starting from a common, inexpensive chiral-pool precursor, L-Malic acid. Key stereocenters and structural motifs were established using Sharpless asymmetric epoxidation, regioselective nucleophilic epoxide opening, Mannich reaction for exo-methylene introduction, and a tandem iodination–zinc elimination cascade. This unified precursor approach offers a streamlined tactical route toward the core macrolide framework.
1. Introduction
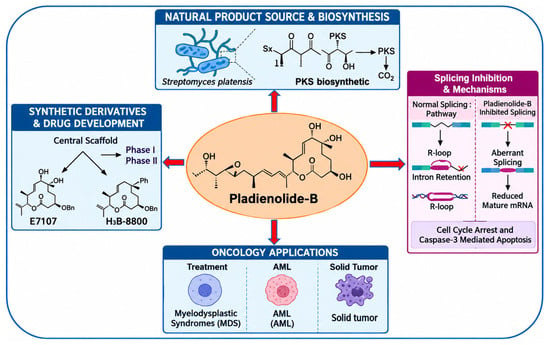
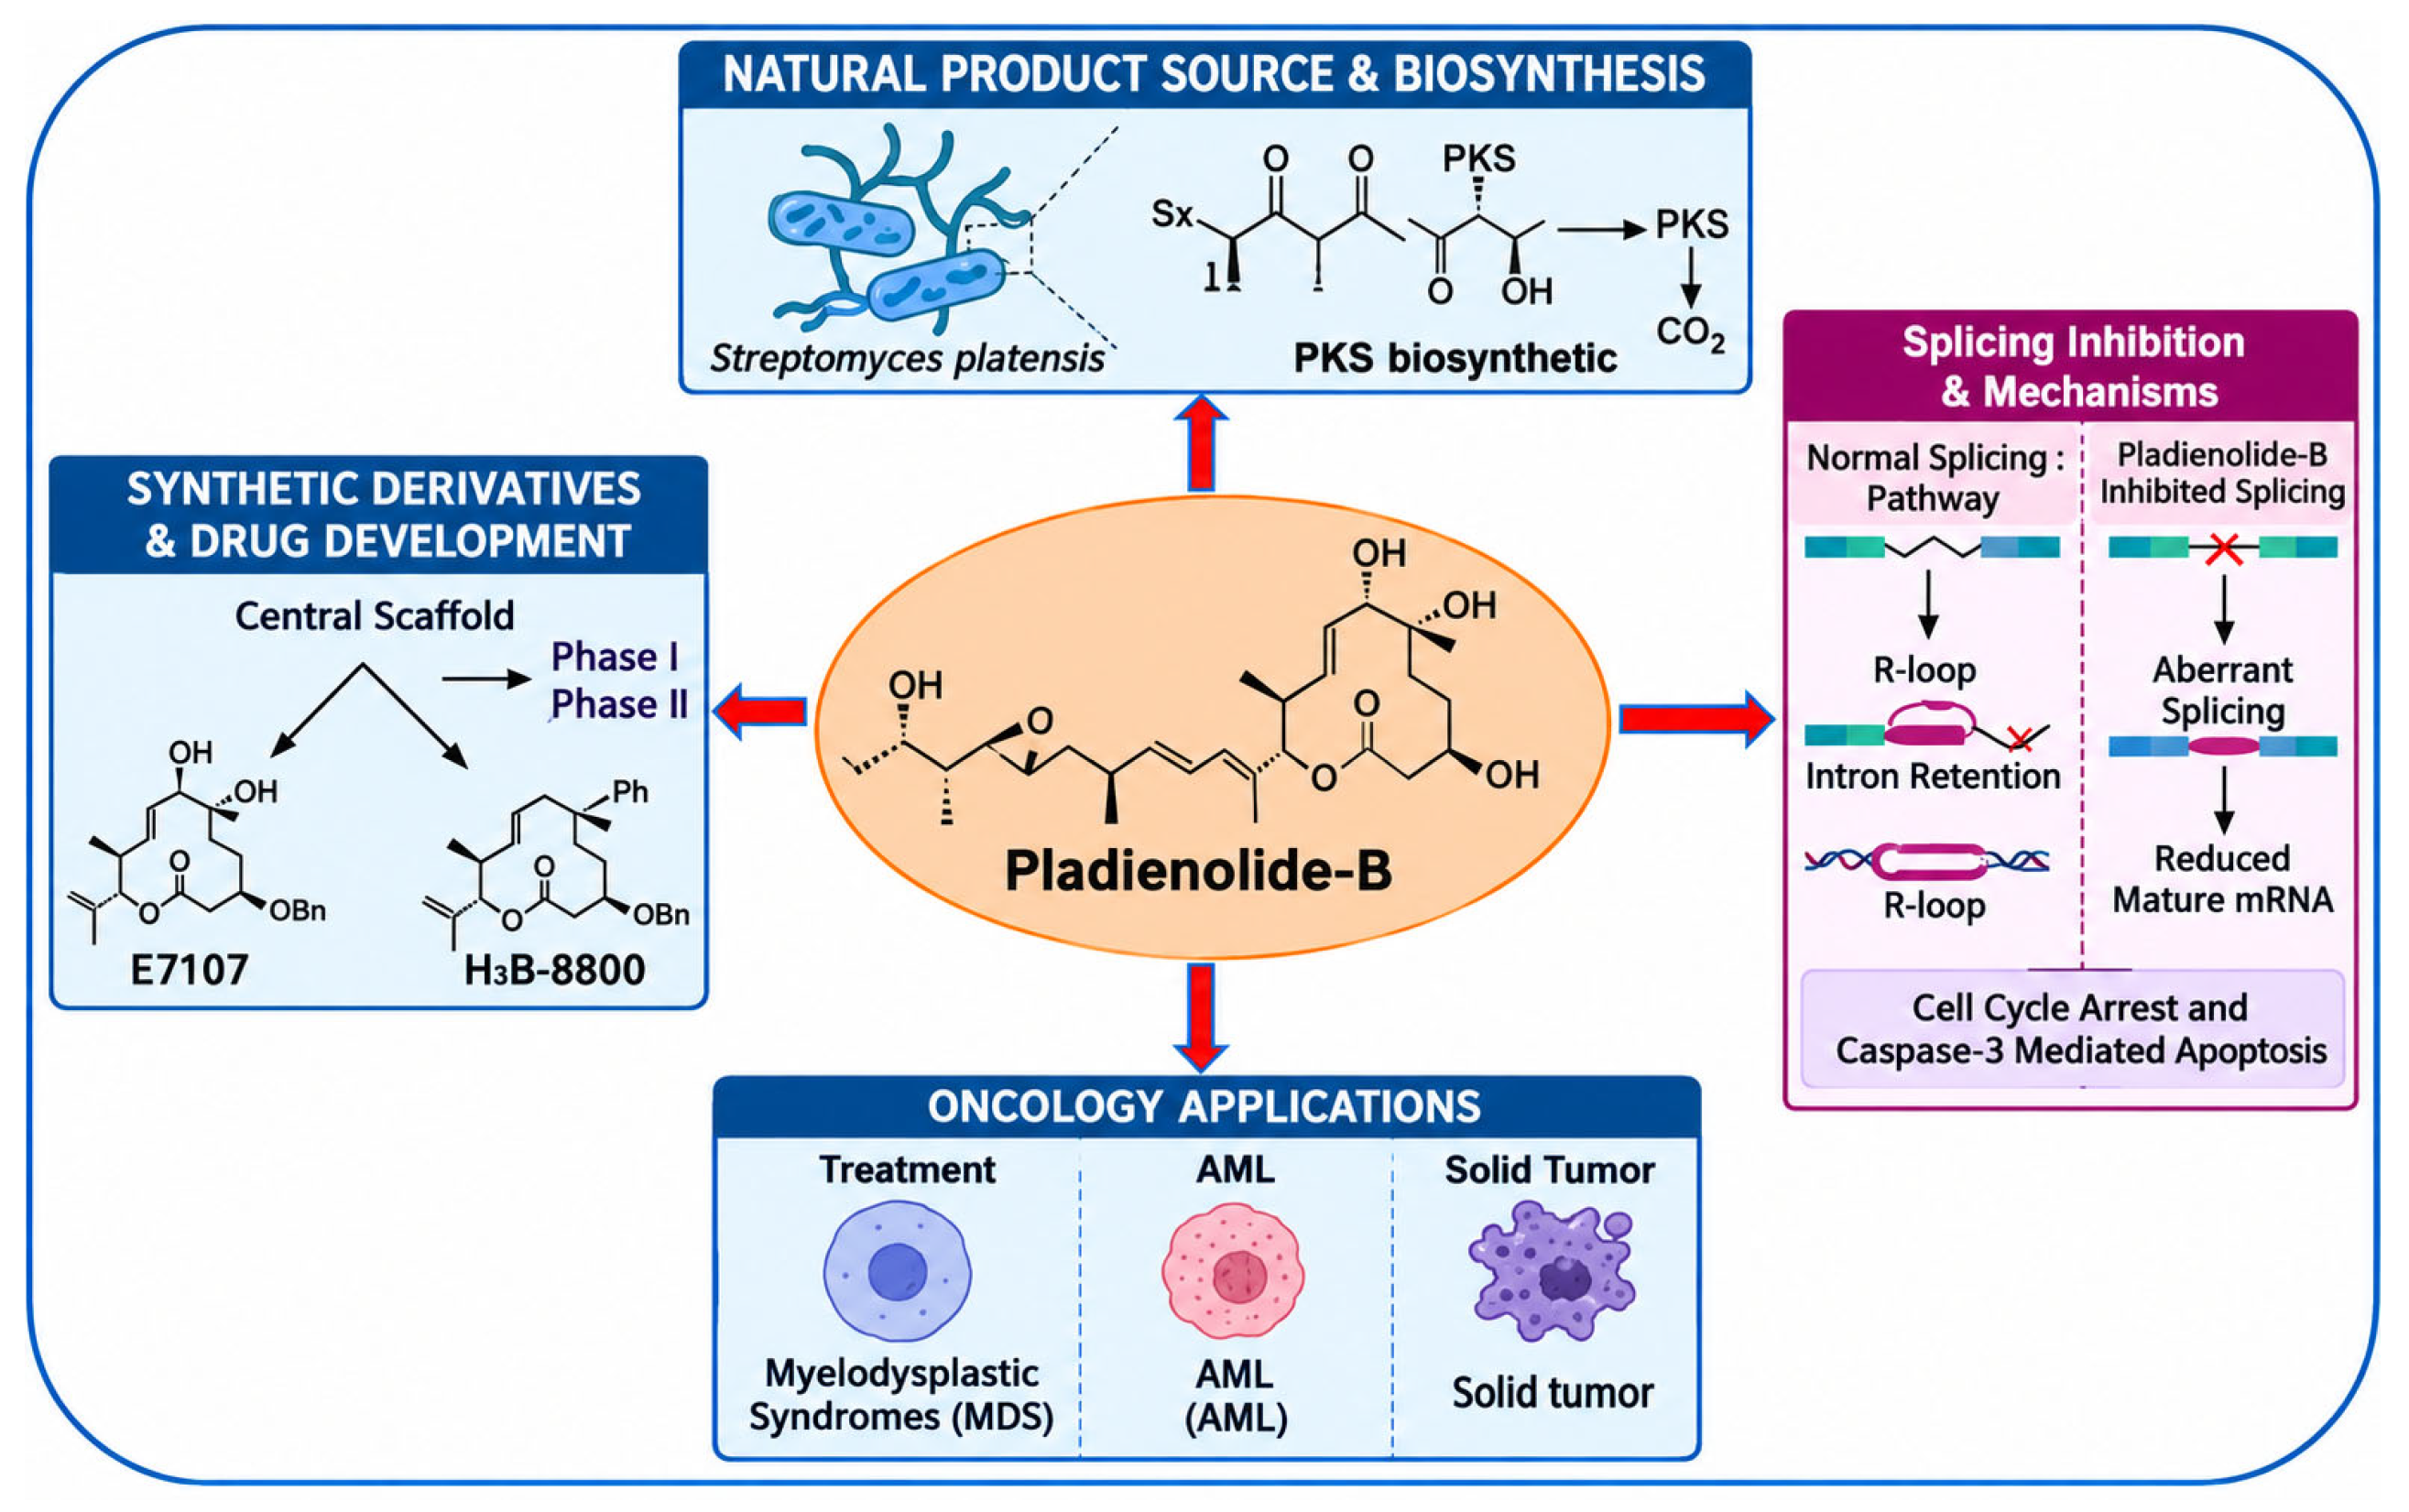
Pladienolides are a family of structurally complex 12-membered macrolides isolated from Streptomyces platensis Mer-11107. Among them, Pladienolide B (1) exhibits potent antiproliferative and antitumor activities and has attracted considerable attention owing to its unique mechanism of action involving the spliceosomal machinery [1,2,3]. Pladienolide B (1) selectively targets the SF3b complex containing SF3B1, thereby disrupting pre-mRNA splicing and inhibiting cancer cell proliferation. This distinctive mode of action has established Pladienolide B (1) as an important lead structure for developing novel anticancer agents and spliceosome-targeting therapeutics [4,5]. More recently, its therapeutic potential has been further demonstrated across different cancer models, including lung adenocarcinoma and hepatocellular carcinoma, where Pladienolide B (1) was shown to suppress tumor-cell proliferation and, in combination with cisplatin, enhance antitumor activity (Figure 1) [6,7,8,9].
Figure 1.
Pladienolide B: Mechanistic Insights and Clinical Applications.
The combination of potent biological activity, a highly functionalized macrolactone framework, multiple stereogenic centres, and an epoxide-containing side chain makes Pladienolide B (1) an attractive target for synthetic studies. Since the isolation and structural elucidation of the pladienolides, several synthetic approaches to Pladienolide B (1) and related analogues have been reported [10,11,12]. The first total synthesis of Pladienolide B (1) and D was reported by the Kotake group, representing a pioneering contribution to the synthesis of these potent antitumor macrolides [13]. Subsequent studies have established diverse strategies for constructing the stereochemically rich macrolide framework, including asymmetric epoxidation, regioselective epoxide opening, olefination, ring-closing metathesis, and stereoselective C–C bond-forming reactions.
These studies have established diverse strategies for constructing the stereochemically rich macrolide framework, including asymmetric epoxidation, regioselective epoxide opening, olefination, ring-closing metathesis, and stereoselective C–C bond-forming reactions. More recently, concise and convergent total syntheses of Pladienolide B (1) and related spliceosome modulators have been developed using catalytic asymmetric C–C bond formation and step-economical synthetic strategies [14,15,16,17].
The C1–C8 and C9–C13 segments constitute important stereochemically defined portions of the Pladienolide B (1). Chiral-pool synthesis provides an attractive strategy for accessing such complex fragments because the inherent stereochemical information of readily available chiral starting materials can be efficiently transferred to the target structures. L-Malic acid, an inexpensive and readily accessible chiral starting material, has been successfully utilized in the synthesis of stereochemically rich intermediates and natural products [18,19,20].
In the present work, we describe a divergent synthesis of the C1–C8 and C9–C13 fragments of Pladienolide B (1) using L-Malic acid as a common chiral-pool starting material. The C1–C8 fragment was constructed through epoxide opening, C–C bond formation, and Sharpless asymmetric epoxidation, followed by conventional functional-group transformations. The C9–C13 fragment was prepared from an L-malic-acid-derived triol through oxidative cleavage, olefination, reduction, and Sharpless asymmetric epoxidation. Thus, two stereochemically defined fragments of the Pladienolide B (1) core were accessed from a common chiral-pool precursor through complementary synthetic sequences.
2. Materials and Methods
All commercially available reagents and solvents, including L-Malic acid, were purchased from Sigma Aldrich (Hyderabad, India), TCI (Hyderabad, India), and Alfa Aesar (Hyderabad, India) and used without further purification unless otherwise noted. All reactions were carried out under an N2 atmosphere in oven-dried glassware using anhydrous solvents (CH2Cl2, THF, CCl4, benzene, or EtOAc), unless otherwise noted. Petroleum ether used had a boiling range of 60–80 °C. The progress of the reactions was monitored by thin-layer chromatography (TLC) using Merck 60 F254 silica gel plates (Hyderabad, India), and visualization was accomplished under UV light 254 nm or by staining with phosphomolybdic acid or KMnO4 solutions. Flash and column chromatography were performed using silica gel (60–120 mesh, Acme Chemical Co. Mumbai, Maharashtra). Reported yields correspond to materials that were homogeneous by chromatographic and spectroscopic (1H NMR, 13C NMR) analysis. Air- and moisture-sensitive reagents were transferred via oven-dried syringes or double-ended cannulas. Concentrating solutions under reduced pressure was performed using a Büchi rotary evaporator (Aditya Scientifics, Hyderabad, Inida). Optical rotations [α]D were measured using a JASCO DIP-370 polarimeter (Aditya Scientifics, Hyderabad, India) at 25 °C. NMR spectra were recorded using Varian Unity 400 MHz or Varian Inova 500 MHz spectrometers (Agilent Technologies, Hyderabad, India). NMR chemical shifts (δ) are given in parts per million (ppm) relative to tetramethyl silane (TMS, δ = 0.00 ppm) as an internal standard using CDCl3 as solvent. Low- and high-resolution mass spectrometric data were obtained using electron impact (70 eV) on an Agilent Technologies LC-MSD spectrometer (Agilent Technologies, Hyderabad, India). The 1H NMR and 13C NMR spectra of compounds 4–8, 11–13, 15–21, 23–27 are provided in the Supplementary Materials.
(R)-1-((4-methoxybenzyl)oxy)hept-6-en-3-ol (11):
A well-stirred suspension of Mg turnings (5.2 g, 216.33 mmol) in diethyl ether (75 mL), allyl chloride (17.6 mL, 216.33 mmol) was added at 0 °C. The Grignard reagent thus generated was stirred until complete formation and then cooled to −40 °C. A solution of the epoxide 9 (15.0 g, 72.11 mmol) in diethyl ether (75 mL) was added slowly to the Grignard solution at −40 °C. The reaction mixture was stirred at −40 °C for 2 h. Upon completion, the reaction mixture was quenched with saturated NH4Cl solution (50 mL) and extracted with EtOAc (2 × 100 mL). The pooled organic layers were washed with water (2 × 50 mL) and brine (2 × 50 mL), dried over anhydrous Na2SO4, and concentrated. The resulting residue was purified by silica gel (60–120 mesh) column chromatography, using 4% EtOAc/pet ether as the eluent, to afford compound 11 as a product (12.6 g, 70% yield). [α]20D = −17.5 (c 0.1, CHCl3); IR (CHCl3): ν 3489, 3487, 3323, 2978, 2922, 1714, 1640, 1609, 1512, 1373, 1246, 1173, 914, 821 C m−1; 1H NMR (300 MHz, CDCl3): δ 7.13 (d, 2H, J = 8.49 Hz), 6.76 (d, 2H, J = 8.49 Hz), 5.72 (m, 1H), 4.94 (m, 2H), 4.36 (s, 2H), 3.72 (s, 3H), 3.51 (m, 3H), 2.7 (bs, 1H), 2.08 (m, 2H), 1.82 (m, 1H), 1.63 (q, 1H), 1.44 (m, 2H); 13C NMR (75 MHz, CDCl3): δ 159.0, 138.4, 131.4, 129.8, 129.1, 114.3, 113.6, 72.7, 70.4, 68.5, 55.0, 36.3, 29.7; HRMS: m/z calculated for C15H22O3Na 273.1388, found: 273.1394.
(R)-1-(((3-(Benzyloxy)hept-6-en-1-yl)oxy)methyl)-4-methoxybenzene (12):
A solution of the compound 11 (12.6 g, 48.0 mmol) in dry THF (10 mL) was added to a cooled suspension of NaH (3.45 g, 144 mmol; 60% w/w dispersion in paraffin oil) in dry THF (15 mL) at 0 °C. The reaction mixture was stirred at 0 °C for 30 min, followed by the dropwise addition of benzyl bromide (6.27 mL, 52.8 mmol) at 0 °C. The reaction mixture was then allowed to warm to room temperature and stirred for 4 h. After completion, the reaction mixture was cooled to 0 °C, and saturated aqueous NH4Cl solution (10 mL) was added dropwise, followed by EtOAc (50 mL). The organic layer was separated, washed successively with water (10 mL) and brine (10 mL), dried over anhydrous Na2SO4, and concentrated under reduced pressure. The resulting residue was purified by column chromatography over silica gel (60–120 mesh) using 5% EtOAc in petroleum ether as the eluent to afford the compound 12 (12.0 g, 74%) as a light yellow color syrup; [α]20D = −37.7 (c 0.1, CHCl3); IR (CHCl3): ν 3065, 3002, 2923, 2854, 1716, 1640, 1512, 1453, 1360, 1246, 1171, 911, 819 cm−1; 1H NMR (500 MHz, CDCl3): δ 7.25 (m, 5H), 7.16 (d, 2H, J = 8.0 Hz), 6.78 (d, 2H, J = 8.0 Hz), 5.75 (m, 1H), 4.93 (m, 2H), 4.44 (q, 2H), 4.35 (q, 2H), 3.77 (s, 3H), 3.51 (m, 2H), 2.11 (m, 2H), 1.78 (m, 2H), 1.61 (m, 2H); 13C NMR (75 MHz, CDCl3): δ 159.1, 138.6, 130.5, 129.3, 128.2, 127.7, 127.4, 114.5, 113.7, 75.6, 72.6, 71.2, 66.5, 55.2, 34.3, 33.4, 29.4; HRMS: m/z calculated for C22H28O3Na 363.1931, found: 363.1944.
(R)-5-(Benzyloxy)-7-((4-methoxybenzyl)oxy)heptan-1-ol (13):
To a solution of cyclohexene (5.4 mL, 52.93 mmol) in THF (10 mL), BH3·DMS (5.0 mL, 52.93 mmol) was added at 0 °C, and the resulting mixture was stirred for 30 min. A solution of the compound 12 (12.0 g, 35.29 mmol) in THF (20 mL) was then added, and the reaction mixture was stirred for an additional 4 h. After completion of the reaction, the reaction mixture was quenched with 2 N NaOH (20 mL), followed by the addition of H2O2 (10 mL). The mixture was extracted with EtOAc (100 mL), and the combined organic extracts were washed with water (2 × 50 mL) and brine (50 mL), dried over anhydrous Na2SO4, and concentrated under reduced pressure. The resulting residue was purified by silica gel column chromatography (60–120 mesh), using 8% EtOAc in petroleum ether as the eluent, to afford the compound 13 (8.84 g, 70%) as liquid; [α]20D = −37.4 (c 0.2, CHCl3); IR (CHCl3): ν 3393, 3360, 2925, 2855, 1712, 1610, 1585, 1456, 1363, 1247, 1173, 1034, 772 cm−1; 1H NMR (500 MHz, CDCl3): δ 7.25 (m, 5H), 7.17 (d, 2H, J = 8.42 Hz), 6.8 (d, 2H, J = 8.42 Hz), 4.45 (q, 2H), 4.36 (t, 2H), 3.78 (s, 3H), 3.58–3.47 (m, 5H), 1.78 (m, 2H), 1.41 (m, 2H), 1.26 (bs, 1H); 13C NMR (75 MHz, CDCl3): δ 159.1, 138.8, 130.5, 129.3, 128.3, 127.7, 127.4, 113.7, 76.1, 72.6, 71.2, 66.5, 62.7, 55.2, 34.3, 33.8, 32.7, 21.3; HRMS: m/z calculated for C22H30O4Na 381.2036, found: 381.2048.
(R)-5-(Benzyloxy)-7-((4-methoxybenzyl)oxy)-2-methyleneheptan-1-ol (6):
Preparation of aldehyde 13a
Dry DMSO (5.26 mL, 74.07 mmol) was added dropwise to a solution of oxalyl chloride (3.17 mL, 37.03 mmol) in dry CH2Cl2 (15 mL) at −78 °C. The reaction mixture was stirred at −78 °C for 30 min, followed by the addition of a solution of the alcohol 13 (8.84 g, 24.69 mmol) in CH2Cl2 (15 mL) over 10 min. The resulting mixture was stirred for 2 h, followed by the dropwise addition of Et3N (20.6 mL, 148.14 mmol). The reaction mixture was then allowed to warm to room temperature and stirred for an additional 30 min. The mixture was diluted with water (10 mL) and CH2Cl2 (50 mL), and the organic layer was separated, washed successively with water (10 mL) and brine (10 mL), dried over anhydrous Na2SO4, and concentrated under reduced pressure to afford the aldehyde 13a (5.0 g, 58% yield).
Preparation of olefin 14
Diethylamine (4.97 mL, 49.11 mmol) and CH2Br2 (16.7 mL, 210 mmol) were mixed and stirred at 55 °C for 1 h 30 min. After cooling the reaction mixture to room temperature, the aldehyde 13a (5.0 g, 14.03 mmol) was introduced, and the mixture was maintained at room temperature for 1 h 30 min. Upon completion, the solvent was evaporated under reduced pressure, and the resulting residue was purified by silica gel column chromatography (60–120 mesh) using 2% EtOAc in petroleum ether as the eluent to furnish the olefin 14 (2.65 g, 45% yield).
Preparation of compound 6
NaBH4 (0.18 g, 5.05 mmol) was added to a solution of the olefin 14 (2.65 g, 7.16 mmol) in CH3OH (10 mL) at 0 °C, and the reaction mixture was stirred at 0 °C until completion. The reaction mixture was then subjected to the appropriate work-up, and the solvent was removed under reduced pressure. The resulting residue was purified by column chromatography over silica gel (60–120 mesh), using 4% EtOAc in petroleum ether as the eluent, to afford the compound 6 (1.15 g, 43%); [α]20D = −87.04 (c 0.2, CHCl3); IR (CHCl3): ν 3440, 3395, 3006, 2859, 2311, 1712, 1609, 1512, 1453, 1360, 1246, 1172, 1029, 820 cm−1; 1H NMR (500 MHz, CDCl3): δ 7.25 (m, 5H), 7.18 (d, 2H, J = 8.3 Hz), 6.81 (d, 2H, J = 8.3 Hz), 4.96 (s, 1H), 4.81 (s, 1H), 4.44 (t, 2H), 4.36 (s, 2H), 3.99 (bs, 1H), 3.78 (s, 3H), 3.54 (m, 3H), 2.12 (m, 2H), 1.8 (m, 2H), 1.68 (q, 2H); 13C NMR (75 MHz, CDCl3): δ 159.1, 148.8, 138.6, 130.4, 129.3, 128.3, 127.8, 127.5, 113.7, 109.3, 75.8, 72.6, 71.2, 66.4, 65.8, 55.2, 34.1, 32.2, 28.4; HRMS: m/z, calculated for C23H30O4Na 393.2144, found 393.2148.
((R)-2-((R)-3-(Benzyloxy)-5-((4-methoxybenzyl)oxy)pentyl)oxiran-2-yl)methanol (15):
Activated 4 Å molecular sieves (0.30 g), Ti(OiPr)4 (0.09 mL, 0.31 mmol), and cumene hydroperoxide (0.97 mL, 6.2 mmol) were added sequentially to a stirred solution of (+)-DIPT (0.13 mL, 0.62 mmol) in CH2Cl2 (15 mL) at −20 °C. The resulting mixture was stirred at −20 °C for 20 min. A solution of the compound 6 (1.15 g, 3.10 mmol) in CH2Cl2 (15 mL) was then added, and the reaction mixture was stirred at −20 °C for 5 h. The reaction was quenched by the addition of 10% KOH solution, prepared by dissolving KOH (3.0 g) in brine (30 mL), and the resulting mixture was stirred for 3 h. The mixture was then filtered through a pad of Celite and washed with EtOAc (20 mL). The organic layer was separated, dried over anhydrous Na2SO4, and concentrated. The resulting residue was purified by silica gel column chromatography using 20% EtOAc in petroleum ether as the eluent, to afford the compound 15 (0.95 g, 80%) as a yellow syrup; [α]20D = −12.2 (c 0.2, CHCl3); IR (CHCl3): ν 3420, 3019, 2985, 1726, 1679, 1514, 1215, 1033, 928 cm−1; 1H NMR (500 MHz, CDCl3): δ 7.27 (m, 7H, ArH), 6.87 (d, 2H, J = 8.3 Hz, ArH), 4.47 (m, 2H, ArCH2), 4.40 (s, 2H, ArCH2), 3.80 (s, 3H, OMe), 3.53 (m, 5H, OCH), 2.84 (d, 2H, J = 4.53, epoxy), 2.62 (d, 1H, J = 4.15 Hz, epoxy), 1.69 (m, 6H, CH2); 13C NMR (75 MHz, CDCl3, 295 K): δ 159.06, 148.76, 138.4, 130.3, 129.3, 128.3, 127.8, 127.6, 113.7, 76.3, 72.6, 71.3, 69.0, 66.4, 61.2, 61.1, 55.4, 55.2, 34.1, 33.9, 27.5, 26.2; HRMS: m/z calculated for C23H30O5Na 409.2169, found: 409.2160.
(2R,5R)-5-(Benzyloxy)-7-((4-methoxybenzyl)oxy)-2-methylheptane-1,2-diol (16):
To a cooled suspension of LAH (0.22 g, 6.15 mmol) in dry THF (2 mL) at 0 °C was added a solution of the compound 15 (0.95 g, 2.46 mmol) in dry THF (2 mL). The reaction mixture was stirred at 0 °C and then allowed to warm to room temperature and stirred for 1 h. The reaction mixture was quenched with saturated Na2SO4 solution (2 mL) and stirred for 1 h. The resulting mixture was filtered and washed with EtOAc (2 × 15 mL). The combined organic layers were washed with water (5 mL) and brine (5 mL), dried over anhydrous Na2SO4, and concentrated under reduced pressure. The resulting residue was purified by column chromatography over silica gel, using 20% EtOAc in petroleum ether as the eluent, to furnish the compound 16 (0.66 g, 70%) as a yellow oil; [α]20D = −13.5 (c 0.1, CHCl3); IR (CHCl3): ν 3409, 3063, 3031, 2857, 1701, 1611, 1512, 1456, 1363, 1302, 1248, 1175, 1035, 911 cm−1; 1H NMR (500 MHz, CDCl3): δ 7.28 (m, 7H), 6.87 (d, 2H, J = 8.0 Hz), 4.5 (m, 2H), 4.40 (s, 2H), 3.80 (s, 3H), 3.58 (m, 3H), 3.39 (m, 2H), 1.86 (m, 6H), 1.13 (s, 3H); 13C NMR (75 MHz, CDCl3): δ 159.1, 148.7, 138.3, 130.3, 129.3, 128.3, 127.8, 127.5, 113.7, 76.5, 72.6, 71.2, 69.6, 66.3, 55.2, 34.2, 33.5, 33.4, 27.7, 23.2, 23.0; HRMS: m/z calculated for C23H32O5Na 411.2142, found 411.2145.
(4R,7R)-Ethyl 7-(benzyloxy)-4-hydroxy-9-((4-methoxybenzyl)oxy)-4-methylnon-2-enoate (17):
Preparation of aldehyde 16a
Dry DMSO (0.36 mL, 5.10 mmol) was added dropwise to a solution of oxalyl chloride (0.21 mL, 2.55 mmol) in dry CH2Cl2 (15 mL) at −78 °C. The resulting reaction mixture was stirred at −78 °C for 30 min, followed by the addition of a solution of the alcohol 16 (0.66 g, 1.70 mmol) in CH2Cl2 (15 mL) over 10 min. The reaction mixture was stirred at −78 °C for 2 h, followed by the slow addition of Et3N (1.41 mL, 10.2 mmol). The mixture was then allowed to warm to room temperature and stirred for an additional 30 min. The reaction mixture was diluted with water (10 mL) and CH2Cl2 (50 mL), and the organic layer was separated, washed successively with water (10 mL) and brine (10 mL), dried over anhydrous Na2SO4, and concentrated under reduced pressure to afford the crude aldehyde 16a, which was used directly in the subsequent step without further purification.
Preparation of compound 17
A solution of the crude aldehyde 16a (0.66 g, 1.70 mmol) in benzene (30 mL) was treated with (ethoxycarbonylmethylene)triphenylphosphorane (0.90 g, 2.56 mmol). The reaction mixture was stirred at reflux for 2 h. Upon completion of the reaction, the solvent was removed under vacuum, and the resulting residue was purified by silica gel (60–120 mesh) column chromatography, using 5% EtOAc in petroleum ether as the eluent, to afford the compound 17. (0.46 g, 60%); [α]20D = −22.5, (c 0.2, CHCl3); IR (CHCl3): ν 3452, 2924, 2854, 1715, 1655, 1610, 1513, 1457, 1368, 1251, 1176, 1094, 822 cm−1; 1H NMR (300 MHz, CDCl3): δ 7.28 (m, 7H), 6.88 (m, 3H), 6.02 (d, 1H, J = 15.48 Hz), 4.48 (s, 2H), 4.40 (s, 2H), 4.2 (q, 2H), 3.8 (s, 3H), 3.4 (m, 3H), 1.83–1.61 (m, 6H), 1.28 (s, 6H); 13C NMR (75 MHz, CDCl3): δ 154.6, 130.4, 129.3, 128.3, 127.8, 127.6, 118.5, 113.7, 75.9, 72.6, 72.5, 71.3, 66.4, 55.2, 51.5, 372, 36.8, 33.7, 28.1, 28.0, 27.8; HRMS: m/z calculated for C27H36O6Na 479.2409, found 479.2404.
(4R,7R)-7-(Benzyloxy)-9-((4-methoxybenzyl)oxy)-4-methylnon-2-ene-1,4-diol (18):
The compound 17 (0.46 g, 1.00 mmol) was dissolved in dry CH2Cl2 (20 mL) and cooled to 0 °C. DIBAL-H (1.8 mL, 2.52 mmol; 20% solution in toluene) was added slowly over 15 min, and the reaction mixture was subsequently allowed to reach room temperature and stirred for 4 h. Upon completion, the mixture was cooled to 0 °C and carefully quenched by sequential addition of methanol (1 mL) and sodium potassium tartrate solution (5 mL). The resulting suspension was filtered through a short pad of Celite and rinsed with EtOAc (2 × 50 mL). The combined filtrate was concentrated under reduced pressure, and the residue was purified by silica gel column chromatography (60–120 mesh) using 10% EtOAc in petroleum ether as the eluent to furnish the compound 18 (0.37 g, 90%) as a colorless oil; [α]20D = −31.5 (c 0.2, CHCl3); IR (CHCl3): ν 3420, 2926, 2856, 1713, 1609, 1513, 1455, 1364, 1251, 1220, 1172, 1030, 821 cm−1; 1H NMR (300 MHz, CDCl3): δ 7.71–7.62 (m, 2H), 7.58–7.52 (m, 1H), 7.35–7.27 (m, 2H), 7.25–7.22 (m, 2H), 6.85–6.84 (m, 2H), 5.82–5.68 (m, 2H), 4.53–4.41 (m, 2H), 4.42– 4.38 (m, 2H), 4.13 (d, J = 4.88 Hz, 2H), 3.79 (s, 3H), 3.67–3.57 (m, 1H), 3.57–3.48 (m, 2H), 1.92–1.83 (m, 1H), 1.82–1.74 (m, 1H), 1.72–1.52 (m, 6H), 1.26 (s, 3H); 13C NMR (75 MHz, CDCl3): δ 159.0, 138.4, 138.2, 130.3, 129.3, 128.2, 127.8, 127.7, 127.5, 126.7, 113.6, 76.3, 76.1, 72.5, 72.2, 66.4, 62.9, 55.2, 37.6, 37.4, 34.0, 33.9, 28.0; HRMS: m/z calculated for C25H34O5Na 437.2298, found 437.2308.
(2R,5R)-5-(Benzyloxy)-2-((2S)-3-(hydroxymethyl)oxiran-2-yl)-7-((4-methoxybenzyl)oxy)- heptan-2-ol (19):
A mixture of (−)-DIPT (0.03 g, 0.18 mmol), Ti(OiPr)4 (0.01 mL, 0.04 mmol), and cumene hydroperoxide (0.27 mL, 1.78 mmol) was added sequentially to a stirred suspension of activated 4 Å molecular sieves (0.30 g) in CH2Cl2 (15 mL) at −20 °C. The resulting mixture was maintained at −20 °C for 20 min. A solution of the compound 18 (0.37 g, 0.89 mmol) in CH2Cl2 (15 mL) was then introduced, and stirring was continued at −20 °C for 5 h. The reaction was quenched with 10% KOH solution, prepared by dissolving KOH (3.0 g) in brine (30 mL), and the mixture was stirred for a further 3 h. The reaction mixture was filtered, and the organic layer was separated, dried over anhydrous Na2SO4, and concentrated under reduced pressure. Purification of the resulting residue by column chromatography over silica gel (60–120 mesh), using 20% EtOAc in petroleum ether as the eluent, furnished the compound 19 (0.33 g, 88%) as a yellow syrup; [α]20D = −55.5, (c 0.2, CHCl3); (c 0.1, CHCl3); IR (CHCl3): ν 3424, 2924, 2854, 1713, 1608, 1513, 1460, 1374, 1253, 1219, 1092, 772 cm−1; 1H NMR (500 MHz, CDCl3): δ 7.39–7.21 (m, 7H), 6.87 (d, J = 8.68 Hz, 2H), 4.5 (brs, 2H), 4.4 (m, 2H), 4.41 (bs, 2H), 3.78 (s, 3H), 3.69–3.48 (m, 5H), 3.24–3.16 (m, 1H), 3.00–2.84 (m, 1H), 2.00–1.75 (m, 2H), 1.72–1.55 (m, 4H), 1.27 (s, 3H); 13C NMR (75 MHz, CDCl3): δ 157.7, 138.4, 131.2, 130.8, 130.3, 129.3, 128.7, 128.2, 127.8, 127.5, 113.6, 76.3, 76.2, 72.6, 71.3, 69.0, 66.3, 61.1, 55.4, 34.1, 33.9, 27.4, 26.2; HRMS: m/z calculated for C25H34O6Na 453.2247, found 453.2259.
(3S,4R,7R)-7-(benzyloxy)-9-((4-methoxybenzyl)oxy)-4-methylnon-1-ene-3,4-diol (8):
Preparation of iodo product 19a
The compound 19 (0.33 g, 0.76 mmol) was dissolved in THF and cooled to 0 °C. PPh3 (0.30 g, 1.15 mmol) and imidazole (0.078 g, 1.15 mmol) were then added, followed by the addition of iodine (I2, 0.28 g, 1.15 mmol). The resultant reaction mass was stirred for 30 min at 0 °C and then quenched with aqueous Na2S2O3 solution (2 mL). The mixture was extracted with EtOAc (50 mL), and the organic layer was washed successively with water (50 mL), brine (25 mL) and over anhydrous Na2SO4. The organic layer was concentrated under vacuum to afford the iodo product 19a (0.41 g), which was used directly in the next step without further purification.
Preparation of compound 8
A solution of compound 19a (0.41 g, 7.64 mmol) in CH3OH (6 mL) was treated with Zn (1.99 g, 30.69 mmol) and NaI (1.10 g, 7.67 mmol) at room temperature. The reaction mixture was then heated under reflux for 6 h with stirring. Upon completion, the reaction mixture was filtered through a short pad of Celite and washed with EtOAc (20 mL). The filtrate was concentrated under vacuum, and the resulting residue was purified by column chromatography over silica gel (60–120 mesh), using 5% EtOAc in petroleum ether as the eluent, to afford compound 8 (0.19 g, 61%) as a yellow liquid; [α]20D = −23.5, (c 0.2, CHCl3); (c 0.1, CHCl3); IR (CHCl3): ν 3384, 3059, 2925, 2855, 1712, 1610, 1512, 1438, 1364, 1250, 1176, 1030, 925 cm−1; 1H NMR (300 MHz, CDCl3): δ 7.35 (m, 5H), 7.24 (d, J = 8.69 Hz, 2H), 6.87 (d, J = 8.69 Hz, 2H), 5.87 (m, 1H), 5.30 (m, 1H), 5.12 (m, 1H), 4.49 (bs, 2H), 4.4 (m, 2H), 3.85 (bs, 1H), 3.79 (s, 3H), 3.57 (m, 3H), 1.85 (m, 6H), 1.14 (s, 3H); 13C NMR (75 MHz, CDCl3): δ 159.0, 136.5, 132.0, 131.9, 129.3, 128.5, 128.4, 128.3, 127.8, 127.5, 117.2, 113.7, 77.5, 76.8, 73.9, 72.6, 71.7, 66.5, 55.2, 34.0, 31.6, 27.3, 23.2; HRMS: m/z calculated for C25H34O5Na 437.2298, found 437.2307.
(4R,5S)-4-((R)-3-(Benzyloxy)-5-((4-methoxybenzyl)oxy)pentyl)-4-methyl-2-phenyl-5-vinyl-1,3-dioxolane (20):
The compound 8 (0.19 g, 0.45 mmol) and p-toluenesulfonic acid (p-TSA; 0.078 g, 0.22 mmol) were dissolved in dry CH2Cl2 (25 mL) and cooled to 0 °C. Benzaldehyde dimethyl acetal (0.08 mL, 0.54 mmol) was then added, and the reaction mixture was allowed to warm to room temperature and stirred for 2 h. Upon completion, the reaction mixture was neutralized with Et3N (1 mL), and the solvent was removed under reduced pressure. The resulting residue was purified by column chromatography over silica gel (60–120 mesh), using 5% EtOAc in petroleum ether as the eluent, to afford the compound 20 (0.20 g, 88% yield) as a colourless product. NMR data were recorded for the major isomer; [α]20D = +8.2, (c 0.2, CHCl3); IR (CHCl3): ν 3440, 3395, 3006, 2859, 2311, 1712, 1609, 1512, 1453, 1360, 1246, 1172, 1029, 820 cm−1; 1H NMR (300 MHz, CDCl3): δ 7.50 (m, 2H), 7.29 (m, 10H), 6.86 (m, 2H), 5.89 (m, 2H), 5.39 (m, 1H), 5.28 (m, 1H), 4.45 (m, 4H), 4.22 (m, 1H), 3.79 (s, 3H), 3.57 (m, 3H), 1.76 (m, 4H), 1.34 (s, 3H), 1.23 (m, 2H); HRMS: m/z calculated for C32H38O5Na 525.0715, found 525.0711.
(R)-3-(Benzyloxy)-5-((4R,5S)-4-methyl-2-phenyl-5-vinyl-1,3-dioxolan-4-yl)pentan-1-ol (21):
The compound 20 (0.20 g, 0.39 mmol) was dissolved in a mixture of CH2Cl2/H2O (4 mL/1 mL), and DDQ (0.10 g, 0.43 mmol) was added at 25–30 °C The reaction mixture was stirred at room temperature for 2 h. After completion, the reaction mixture was concentrated under vacuum, and the resulting residue was purified by column chromatography over silica gel (60–120 mesh), using 17% EtOAc in petroleum ether as the eluent, to afford compound 21 (0.1 g, 72%) as a colorless oil; [α]20D = +8.26, (c 0.2, CHCl3); IR (CHCl3): ν 3422, 3032, 2924, 2854, 1719, 1454, 1378, 1273, 1219, 1176, 1093, 1027, 933, 772 cm−1; 1H NMR (300 MHz, CDCl3) δ 7.48 (m, 2H), 7.34 (m, 8H), 5.91 (m, 2H), 5.43 (m, 1H), 4.60 (m, 1H), 4.39 (m, 1H), 4.25 (m, 1H), 3.69 (m, 3H), 2.28 (brs, 1H), 1.79 (m, 6H), 1.34 (s, 3H); 13C NMR (75 MHz, CDCl3): δ 139.3, 138.2, 133.1, 129.5, 129.0, 128.9, 128.4, 128.2, 127.8, 126.7, 126.5, 126.2, 125.9, 118.9, 102.0, 87.7, 86.0, 83.5, 78.4, 78.2, 77.9, 70.8, 60.6, 35.8, 35.6, 28.0, 26.8, 26.2, 21.8; HRMS: m/z calculated for C24H34O4N 400.0015, found 400.0021.
(R)-3-(Benzyloxy)-5-((4R,5S)-4-methyl-2-phenyl-5-vinyl-1,3-dioxolan-4-yl)pentanoicacid (4):
The alcohol 21 (0.10 g, 0.26 mmol) was dissolved in a mixture of CH2Cl2/H2O (20 mL/1 mL) and cooled to 0 °C. TEMPO (0.012 g, 0.07 mmol) was then added, followed by BAIB (0.25 g, 0.78 mmol). The reaction mixture was allowed to warm to room temperature and stirred for 1 h. Upon completion, the reaction mixture was quenched with aqueous sodium thiosulfate (Na2S2O3) solution (50 mL). The organic layer was separated, washed successively with water (50 mL) and brine (25 mL), dried over anhydrous Na2SO4, and concentrated under reduced pressure. The resulting residue was purified by column chromatography over silica gel (60–120 mesh), using 70% EtOAc in petroleum ether as the eluent, to afford the compound 4. (0.06 g, 60%) as a light-yellow oil. [α]20D = +15.4 (c 0.2, CHCl3); IR (CHCl3): ν 2925, 2854, 1730, 1459, 1380, 1219, 1091, 1068, 7726 cm−1; 1H NMR (300 MHz, CDCl3): δ 9.73 (s, 1H), 7.48 (m, 2H), 7.34 (m, 8H), 5.91 (m, 2H), 5.43 (m, 2H), 4.60 (m, 1H), 4.39 (m, 1H), 4.25 (m, 1H), 3.69 (m, 1H), 2.28 (m, 2H), 1.79 (m, 4H), 1.34 (s, 3H). 13C NMR (75 MHz, CDCl3) δ 159.1, 132.2, 130.0, 129.6, 128.6, 123.5, 113.6, 71.7, 68.5, 65.8, 55.1; HRMS: m/z calculated for C24H27O5 395.0019 found 395.0011.
(R)-Ethyl 5-hydroxy-2,4-dimethylpent-2-enoate (23):
Preparation of the aldehyde 22a
Compound 22 (5.0 g, 41.66 mmol) was dissolved in a 5:1 mixture of acetone/H2O (30 mL) and cooled to 0 °C. NaIO4 (10.7 g, 50.0 mmol) was then added, and the reaction mixture was allowed to warm to room temperature and stirred for 1 h. Upon completion, the solvent was removed under reduced pressure, and the residue was extracted with CHCl3 (2 × 25 mL). The combined organic extracts were concentrated under reduced pressure to afford the aldehyde 22a (3.66 g), which was carried forward to the next step without further purification.
Preparation of the compound 23
The Wittig ylide (20.17 g, 57.88 mmol) was dissolved in toluene (10 mL), and a solution of the aldehyde 22a (3.96 g, 48.24 mmol) in toluene (10 mL) was added. The resulting mixture was heated under reflux for 2 h with stirring. Upon completion, the solvent was removed under reduced pressure, and the residue was purified by column chromatography over silica gel (60–120 mesh), using 15% EtOAc in petroleum ether as the eluent, to afford the compound 23 (5.07 g, 71%) as a colorless oil; [α]20D = −13.6 (c 0.2, CHCl3); IR (CHCl3): ν 3456, 3017, 2978, 2933, 2874, 1708, 1648, 1451, 1390, 1263, 1214, 1131, 1091, 930, 750 cm−1; 1H NMR (300 MHz, CDCl3): δ 6.50 (d, 1H, J = 11.33), 4.18 (q, 2H), 3.51 (m, 2H, OCH), 2.74 (m, 1H), 1.88 (s, 3H), 1.62 (bs, 1H), 1.31 (t, 3H), 1.03 (d, 3H); 13C NMR (75 MHz, CDCl3, 295 K) δ 168.1, 143.8, 129.1, 67.0, 60.6, 36.1, 16.0, 14.2, 12.7; HRMS: m/z calculated for C9H16O3Na 199.0992, found 199.0995.
(R)-Ethyl 5-((tert-butyldiphenylsilyl)oxy)-2,4-dimethylpent-2-enoate (24):
To a solution of ester 23 (5.07 g, 29.47 mmol) in CH2Cl2 (25 mL) at 0 °C, imidazole (6.01 g, 88.43 mmol) and TBDPSCl (9.68 g, 35.36 mmol) were added sequentially and the reaction mixture was stirred at 0 °C for 1 h. Upon completion of the reaction, the solvent was removed under vacuum, and the resulting residue was purified by silica gel (60–120 mesh) column chromatography, using 3% EtOAc in petroleum ether as the eluent, to afford compound 24 (8.45 g, 70%); [α]20D = −66.2 (c 0.1, CHCl3); IR (CHCl3): ν 3070, 3050, 2959, 2931, 2896, 2858, 1711, 1651, 1588, 1469, 1427, 1365, 1266, 1234, 1109, 1083, 803 cm−1; 1H NMR (300 MHz, CDCl3): δ 7.62 (m, 4H), 7.35 (m, 6H), 6.53 (d, 1H, J = 10.00 Hz), 4.17 (q, 2H), 3.52 (d, 2H), 2.71 (m, 1H), 1.78 (d, 3H), 1.29 (t, 3H), 1.04 (s, 9H); 13C NMR (75 MHz, CDCl3, 295 K): δ 168.2, 144.4, 135.6, 133.6, 129.6, 127.9, 127.6, 67.7, 60.3, 36.1, 26.8, 19.2, 16.3, 14.3, 12.6, HRMS: m/z calculated for C25H34O3SiNa 433.1029 found 433.1024.
(R)-5-(tert-Butyldiphenylsilyloxy)-2,4-dimethylpent-2-en-1-ol (7):
Compound 24 (8.45 g, 20.6 mmol) was dissolved in dry CH2Cl2 (20 mL) and cooled to 0 °C. DIBAL-H (21.3 mL, 30.91 mmol; 20% solution in toluene) was added dropwise over 15 min, and the reaction mixture was subsequently allowed to warm to room temperature and stirred for 4 h. The mixture was then cooled to 0 °C and carefully quenched by sequential addition of methanol (1 mL) and sodium potassium tartrate solution (5 mL). The resulting mixture was filtered through a short pad of Celite and washed with EtOAc (2 × 50 mL). The filtrate was concentrated under reduced pressure, and the residue was purified by column chromatography over silica gel (60–120 mesh), using 10% EtOAc in petroleum ether as the eluent, to afford the compound 7 (6.0 g, 72%) as a colorless oil; [α]20D = −57.2 (c 0.2, CHCl3); IR (CHCl3): ν 3440, 3395, 3019, 2958, 2930, 2858, 1468, 1427, 1214, 1110, 1082, 1007, 931, 741 cm−1; 1H NMR (500 MHz, CDCl3): δ 7.62 (m, 4H), 7.36 (m, 6H), 5.2 (m, 1H), 3.9 (m, 2H), 3.46 (m, 2H), 2.6 (m, 1H), 1.6 (d, 2H), 1.32 (bs, 1H), 1.05 (s, 9H), 0.99 (d); 13C NMR (75 MHz, CDCl3): δ 135.6, 133.9, 129.5, 128.9, 127.5, 68.8, 68.4, 35.0, 26.8, 19.2, 17.2, 13.8; HRMS: m/z calculated for C23H32O2SiNa 391.0105 found 391.0109.
((2S,3S)-3-((S)-1-(tert-Butyldiphenylsilyloxy)propan-2-yl)-2-methyloxiran-2-yl)methanol (25):
A stirred solution of (+)-DIPT (0.78 g, 3.35 mmol) in CH2Cl2 (15 mL) containing activated 4 Å molecular sieves (0.30 g) was cooled to −20 °C. Ti(OiPr)4 (0.47 g, 1.67 mmol) and cumene hydroperoxide (5.14 g, 33.5 mmol) were then added sequentially, and the resulting mixture was stirred at −20 °C for 20 min. A solution of the compound 7 (6.0 g, 16.75 mmol) in CH2Cl2 (15 mL) was subsequently added, and the reaction mixture was maintained at −20 °C for 5 h. The reaction was quenched with 10% KOH solution, prepared by dissolving KOH (3.0 g) in brine (30 mL), and the mixture was stirred for an additional 3 h. The resulting mixture was filtered, and the organic layer was separated, dried over anhydrous Na2SO4, and concentrated under reduced pressure. The residue was purified by column chromatography over silica gel (60–120 mesh), using 20% EtOAc in petroleum ether as the eluent, to afford the compound 25 (4.0 g, 62%) as a yellow oil; [α]20D = −87.04 (c 0.1, CHCl3); IR (CHCl3): ν 3439, 3397, 3070, 2958, 2930, 2857 1726, 1589, 1469, 1427, 1388, 1216, 1187 1107, 1029, 889, 740 cm−1; 1H NMR (500 MHz, CDCl3): δ 7.57 (m, 5H), 7.29 (m, 5H), 3.66 (m, 2H), 3.50 m, 2H), 2.79 (m, 1H), 1.5 (m, 1H), 1.18 (bs, 3H), 1.04 (s, 9H), 0.97 (d, 6H); 13C NMR (75 MHz, CDCl3): δ 135.6, 129.5, 127.6, 66.3, 65.4, 61.3, 35.4, 26.8, 14.1, 13.4; HRMS: m/z calculated for C23H32O3NaSi 407.2013, found 407.2017.
(3S,4S)-5-(tert-Butyldiphenylsilyloxy)-2,4-dimethylpent-1-en-3-ol (26):
Preparation of the iodo compound 25a
Compound 25 (4.0 g, 10.41 mmol) was dissolved in THF, followed by the sequential addition of PPh3 (4.0 g, 15.62 mmol), imidazole (2.12 g, 31.23 mmol), and iodine (I2, 3.93 g, 15.61 mmol). The reaction mixture was stirred at room temperature for 30 min and then quenched with saturated aqueous NaOH solution (5 mL). The mixture was extracted with EtOAc (100 mL), and the organic layer was washed successively with aqueous sodium thiosulfate (hypo) solution (50 mL), water (50 mL), and brine (25 mL). The organic layer was dried over anhydrous Na2SO4 and concentrated under reduced pressure to afford the compound 25a (5.15 g), which was carried forward to the next step without further purification.
Preparation of the compound 26
A solution of compound 25a (5.15 g, 10.42 mmol) in CH3OH (25 mL) was treated with Zn (2.0 g, 31.27 mmol) and NaI (2.32 g, 15.63 mmol). The reaction mixture was heated under reflux and stirred for 6 h. Upon completion, the mixture was filtered through a short pad of Celite and washed with EtOAc (20 mL). The filtrate was concentrated under vacuum, and the resulting residue was purified by column chromatography over silica gel (60–120 mesh), using 5% EtOAc in petroleum ether as the eluent, to afford compound 26 (2.37 g, 62%) as a yellow oil; [α]20D = −37.4 (c 0.2, CHCl3); IR (CHCl3): ν 3439, 3397, 3070, 2958, 2930, 2857 1726, 1632, 1620, 1589, 1490, 1469, 1427, 1388, 1216, 1187 1107, 1029, 889, 740 cm−1; 1H NMR (300 MHz, CDCl3): δ 7.65 (m, 5H), 7.38 (m, 5H), 4.93 (m, 2H), 4.27 (bs, 1H), 3.95 (d, 1H, J = 8.12 Hz), 3.72 (m, 2H), 1.85 (m, 1H), 1.68 (s, 3H), 1.27 (s, 3H), 1.07 (s, 9H), 0.8 (d, 3H); 13C NMR (75 MHz, CDCl3): δ 159.1, 138.3, 130.3, 129.3, 128.3, 127.8, 127.5, 113.7, 76.5, 72.6, 72.4, 71.2, 69.6, 66.3, 55.2, 33.9, 33.5, 27.7, 23.2, 23.0; HRMS: m/z calculated for C23H32O2NaSi 391.0015, found 391.0019.
(3S,4S)-5-(tert-Butyldiphenylsilyloxy)-2,4-dimethylpent-1-en-3-yl benzoate (27):
The compound 26 (2.37 g, 6.44 mmol) was dissolved in CH2Cl2 and cooled to 0 °C. Et3N (1.8 mL, 12.88 mmol) and benzoyl chloride (BzCl, 0.9 mL, 6.44 mmol) were then added sequentially, followed by a catalytic amount of DMAP. The reaction mixture was allowed to warm to room temperature and stirred for 3 h. Upon completion, the solvent was removed under reduced pressure, and the resulting residue was purified by column chromatography over silica gel (60–120 mesh), using 15% EtOAc in petroleum ether as the eluent, to afford the compound 27 (1.97 g, 65%) as syrup; [α]20D = +22.5 (c 0.2, CHCl3); IR (CHCl3): ν 2958, 2930, 2857 1726, 1632, 1620, 1589, 1490, 1469, 1427, 1388, 1216, 1187 1107, 1029, 889, 740 cm−1; 1H NMR (300 MHz, CDCl3): δ 8.05 (d, 2H, J = 7.17 Hz), 7.72 (d, 3H, J = 7.17 Hz), 7.74 (m, 10H), 5.01 (m, 2H), 4.41 (dd, 1H), 4.21 (m, 1H), 2.33 (m, 1H), 2.07 (t, 3H), 1.78 (d, 3H), 1.05 (s, 9H, CH3); 13C NMR (75 MHz, CDCl3): δ 165.4, 141.5, 135.5, 133.5, 132.6, 130.5, 129.5, 129.4, 128.2, 127.5, 127.4, 114.9, 79.2, 65.3, 64.6, 53.3, 37.4, 29.6, 26.7, 19.2, 17.8, 13.9; HRMS: m/z calculated for C30H36O3NaSi 495.0011, found 495.0015.
(3S,4S)-5-hydroxy-2,4-dimethylpent-1-en-3-yl benzoate (5):
The compound 27 (1.97 g, 4.17 mmol) was dissolved in anhydrous THF (10 mL) and cooled to 0 °C. TBAF (4.2 mL, 4.17 mmol; 1.0 M solution in THF) was then added, and the reaction mixture was allowed to warm to room temperature and stirred for 3 h. Upon completion, the reaction mixture was diluted with water (25 mL) and extracted with EtOAc (2 × 25 mL). The combined organic extracts were washed successively with water (20 mL) and brine (20 mL), dried over anhydrous Na2SO4, and concentrated under reduced pressure. The resulting residue was purified by column chromatography over silica gel (60–120 mesh), using 20% EtOAc in petroleum ether as the eluent, to afford the compound 5 (0.58 g, 60%) as a colorless oil; [α]D25 +12.5 (c 0.2, CHCl3); IR (neat): 3416, 2935, 1711, 1608, 1514, 1441, 1252, 1101, 1034, 916, 821 cm−1; 1H NMR (300 MHz, CDCl3): δ 8.05 (d, 2H, J = 6.29 Hz), 7.58 (t, 1H, J = 7.32 Hz), 7.46 (t, 2H), 5.37 (d, J = 9.0 Hz, 1H), 5.07 (s, 1H), 5.01 (s, 1H), 3.65 (m, 2H), 2.09 (m, 2H), 1.8 (s, 3H) 1.01 (d, 3H); 13C NMR (75 MHz, CDCl3): δ 166.3, 141.6, 134.7, 133.1, 130.0, 129.6, 128.4, 127.6, 115.1, 79.9, 63.9, 37.2, 26.5, 17.8, 13.8; HRMS: m/z calculated for C14H18O3Na 257.1148, found 257.1151.
3. Results
3.1. Retrosynthesis
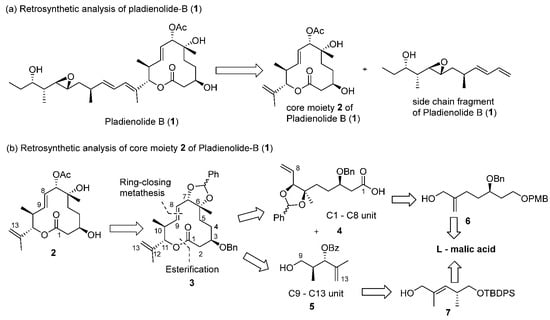
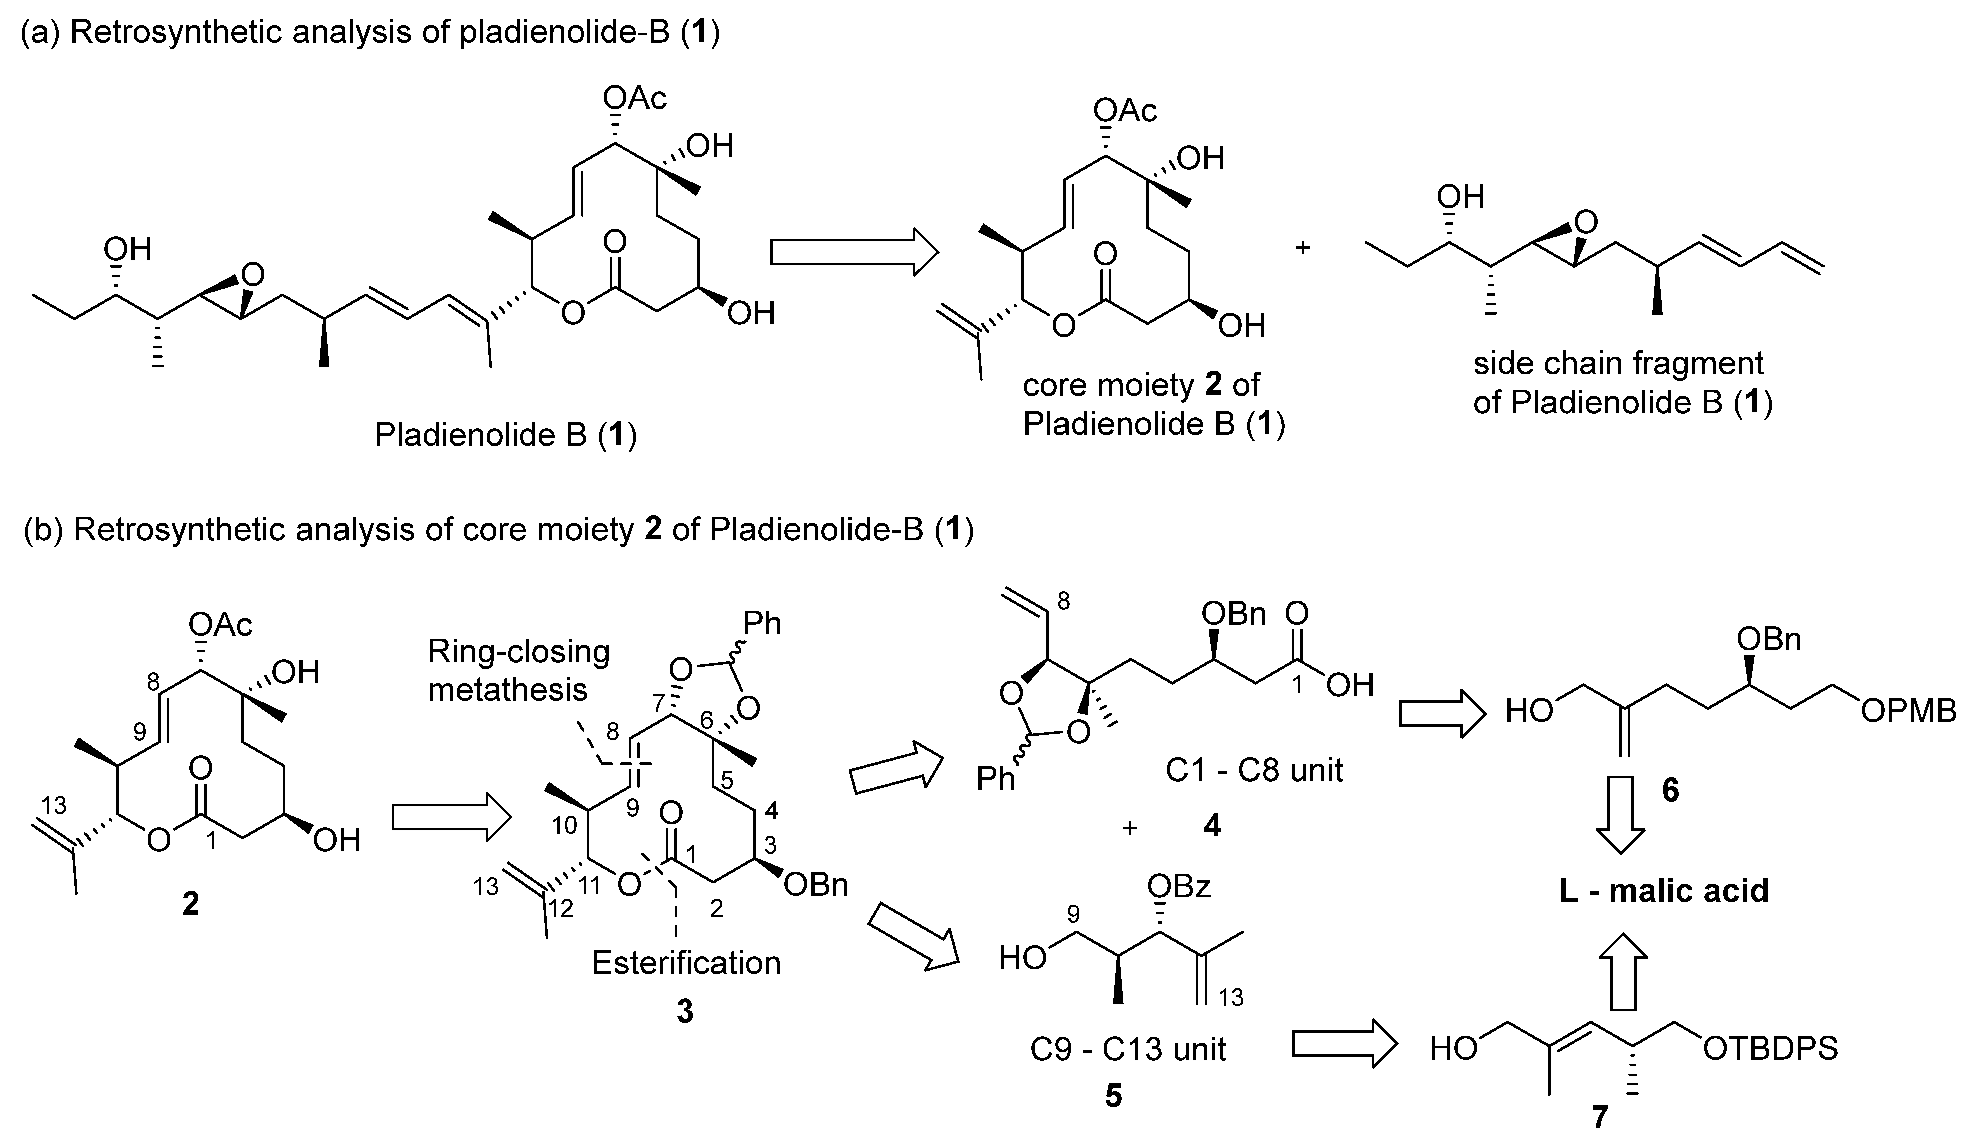
The synthesis of Pladienolide B (1) was envisaged from its core moiety 2 and the corresponding side-chain fragment, as shown in Scheme 1a. The macrolide core was designed to be assembled through esterification of the C1–C8 acid fragment 4 with the C9–C13 alcohol fragment 5, followed by ring-closing olefin metathesis (RCM). Retrosynthetic analysis of 2 identified a convergent disconnection into two polyketide subunits, which were traced back to intermediates 6 and 7, respectively (Scheme 1b) [13]. To minimize the use of multiple non-overlapping chiral starting materials and improve the overall efficiency and cost-effectiveness of the synthesis, both intermediates were derived from the common chiral-pool precursor L-Malic acid 10.
Scheme 1.
Retrosynthetic analysis of Pladienolide B (1).
3.2. Synthesis of C1–C8 Fragment 4 of Pladienolide B (1)
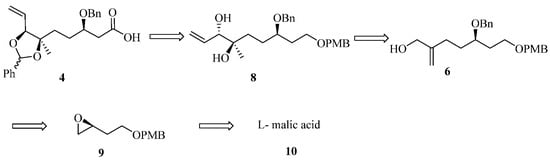
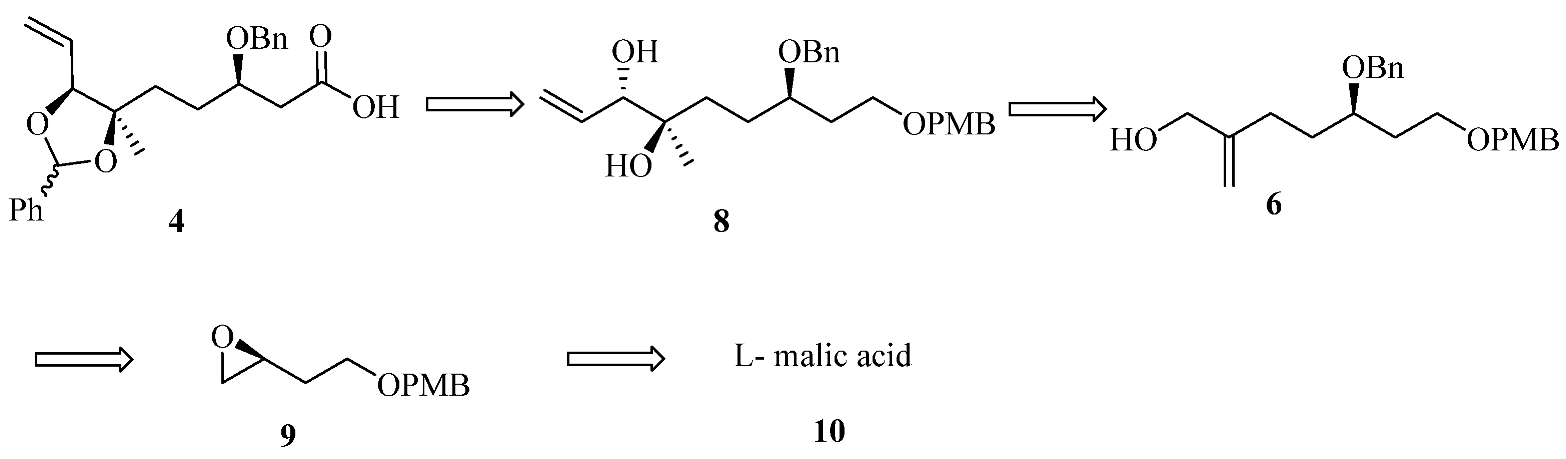
The retrosynthetic plan for the C1–C8 acid fragment 4 is depicted in Scheme 2. Target fragment 4 was envisioned to be accessible from the 1,2-diol 8, which in turn could be constructed from the allylic alcohol 6 via a sequence involving Sharpless asymmetric epoxidation, regioselective epoxide opening, and functional group transformations. The allylic alcohol 6 could be derived through short, sequential manipulations of known epoxide 9 [21], prepared from L-Malic acid 10.
Scheme 2.
Retrosynthetic analysis of acid fragment 4.
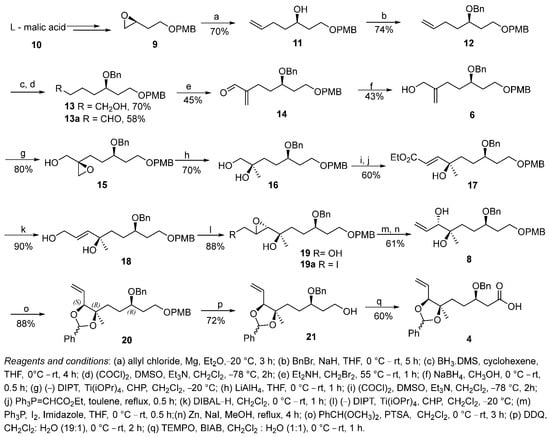
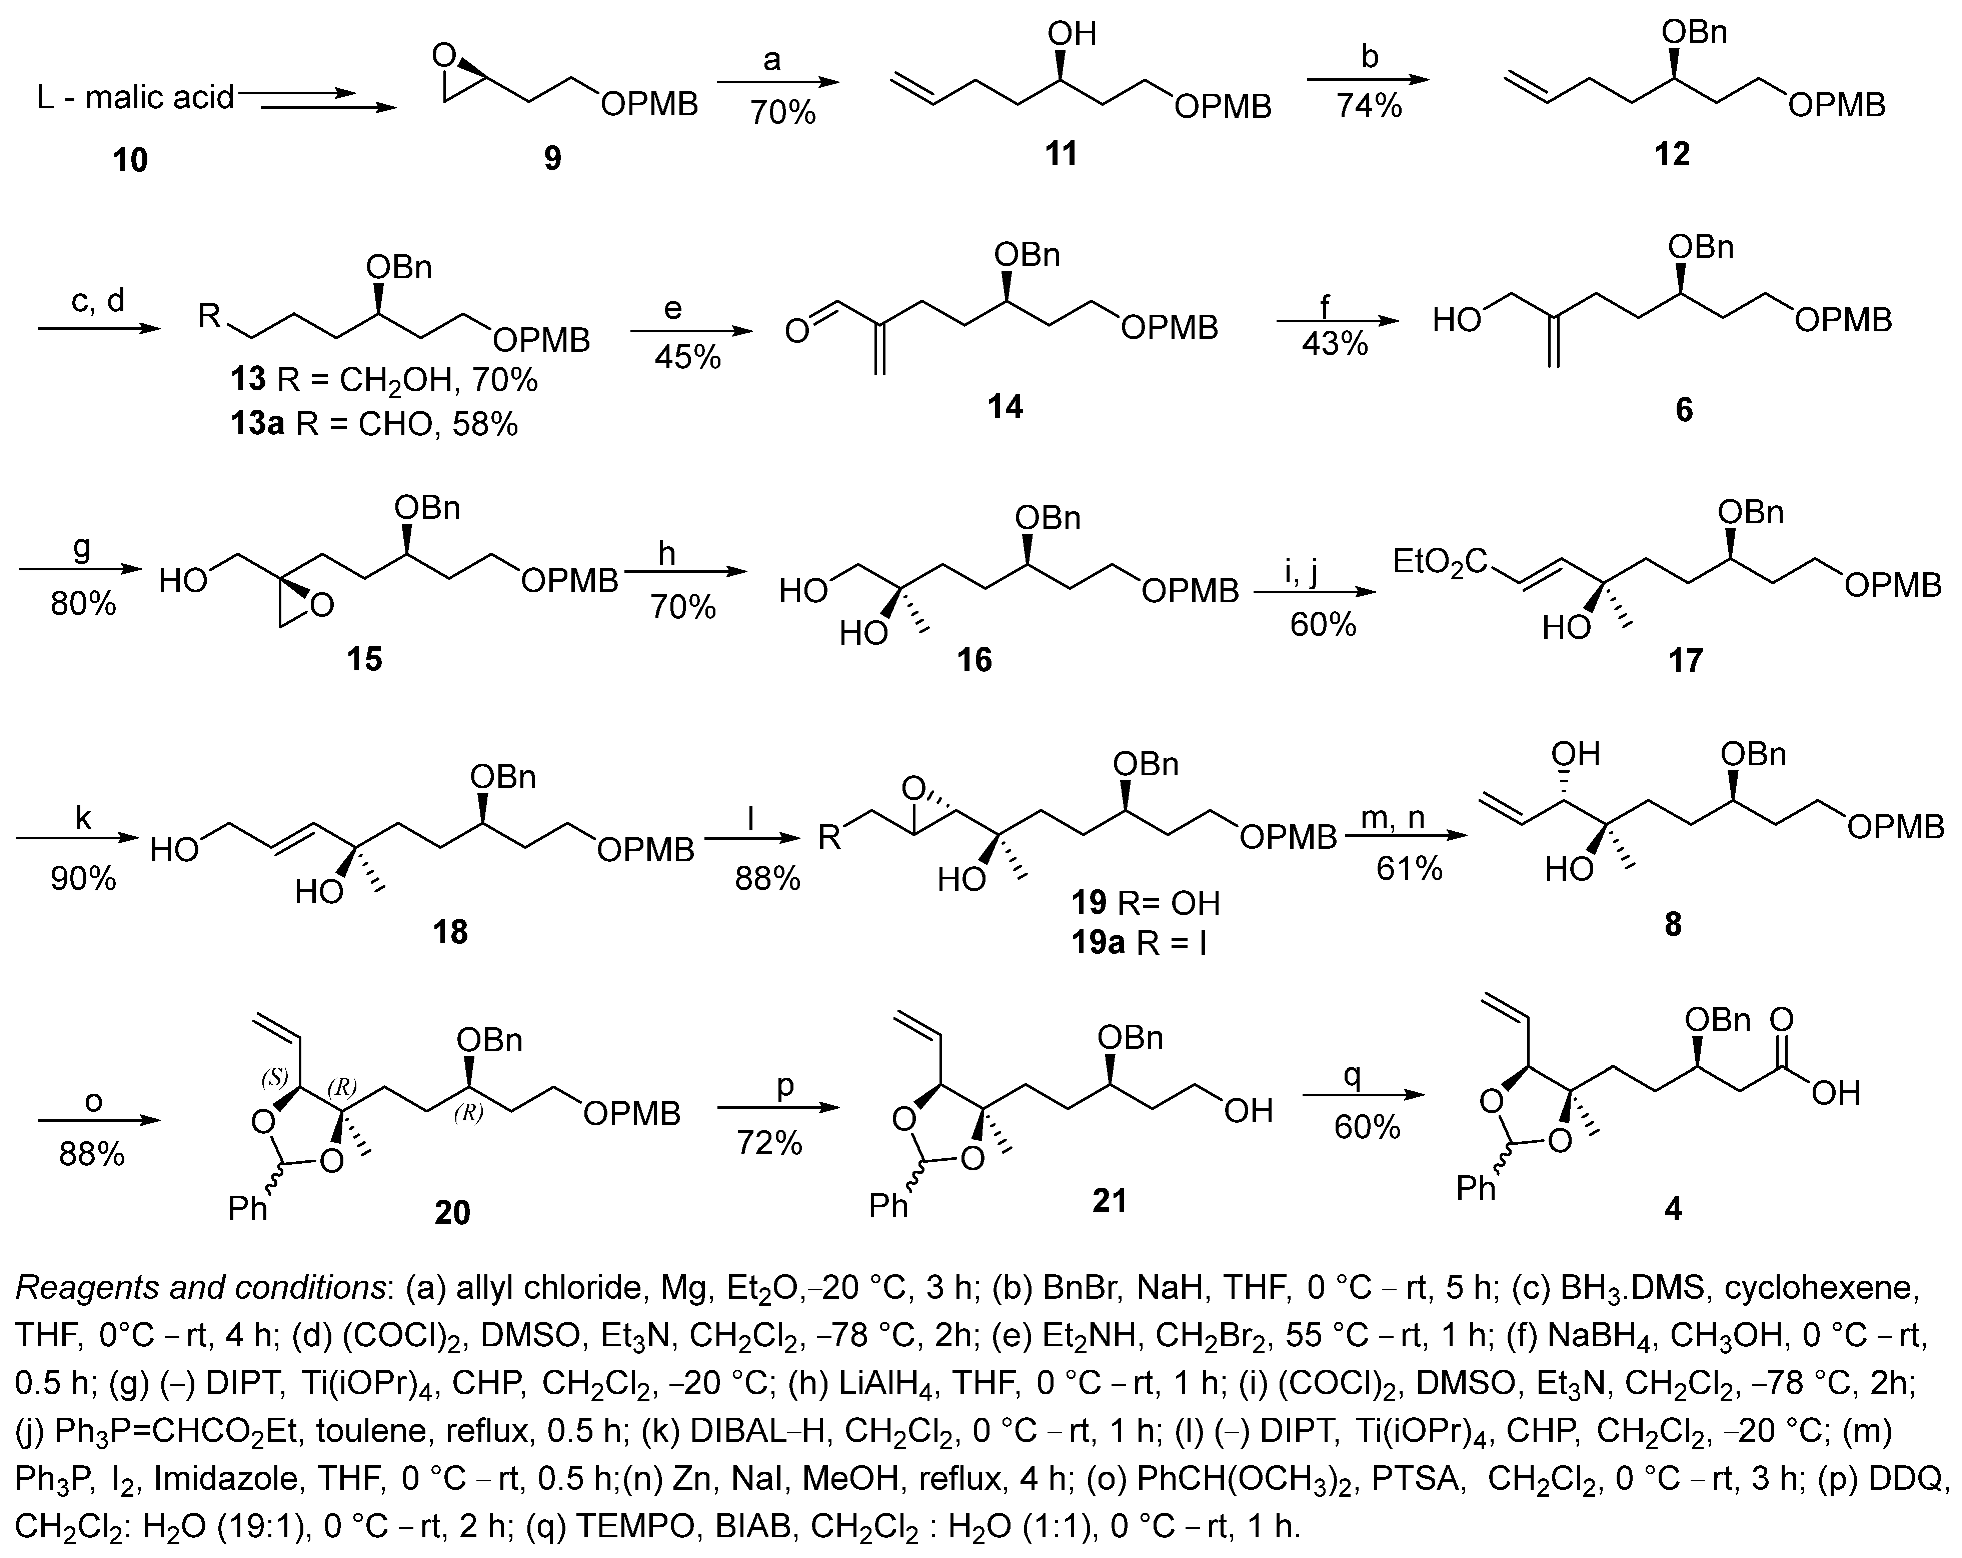
The forward synthesis commenced with the regioselective ring opening of the epoxide 9 by Grignard reaction with allyl magnesium chloride in ether at −20 °C for 3 h, delivering secondary alcohol 11 in 80% yield (Scheme 3). O-Benzylation of the alcohol 11 using benzyl bromide and sodium hydride in anhydrous THF at 0 °C for 5 h afforded the corresponding benzyl ether 12. Selective hydroboration–oxidation of the terminal alkene in 12 with BH3.DMS and cyclohexene in THF at 0 °C for 4 h furnished the primary alcohol 13. Subsequent Swern oxidation of 13a yielded aldehyde 13b, which was then subjected to a Mannich methylene insertion reaction using a preheated mixture of diethylaminem diethylamine and dibromo methane at r.t. for 3 h to afford the α,β-unsaturated aldehyde 14 (enal). Chemo-selective reduction of enal 14 using sodium borohydride in methanol at 0 °C for 0.5 h smoothly afforded the allylic alcohol 6. Sharpless asymmetric epoxidation of the allylic alcohol 6 using L-(+)-diethyl tartrate, Ti(iOPr)4, and tert-butyl hydroperoxide in dichloromethane at −20 °C successfully proceeded the required epoxy stereocenters in the compound 15. Regioselective reduction of epoxy alcohol 15 with lithium aluminum hydride (LiAlH4) in THF at 0 °C for 1 h afforded the 1,2-diol 16. Subsequent Swern oxidation of 16, followed by Wittig homologation using methyl (triphenylphosphoranylidene)-acetate, afforded the corresponding α,β-unsaturated ester 17.
Scheme 3.
Synthesis of C1–C8 fragment 4 of Pladienolide B (1).
Reduction of the ester 17 using DIBAL-H in dichloromethane at 0 °C for 1 h afforded the corresponding allylic alcohol 18. A second Sharpless asymmetric epoxidation of the allylic alcohol 18 under identical kinetic resolution conditions delivered the epoxy alcohol 19. Treatment of the 19 under Appel-type iodination conditions led to the iodomethyl intermediate 19a. Tandem zinc-promoted reductive elimination of the compound 19a using zinc powder and sodium iodide in refluxing methanol for 4 h cleanly installed the exo-methylene motif, yielding the diol 8.
Protection of the diol 8 with benzaldehyde dimethyl acetal using catalytic p-toluene sulfonic acid (pTSA) in dichloromethane at 0 °C for 3 h afforded the benzylidene acetal 20. Oxidative deprotection of the secondary benzyl ether in the compound 20 using 2,3-dichloro-5,6-dicyano-1,4-benzoquinone (DDQ) in DCM/H2O mixture (19:1) provided the primary alcohol 21. Finally, TEMPO/BAIB-mediated oxidation of the primary alcohol 21 in DCM/H2O mixture (19:1) at room temperature for 1 h successfully furnished the target C1–C8 acid fragment 4.
3.3. Synthesis of the C9–C13 Alcohol Fragment 5 of Pladienolide B (1)
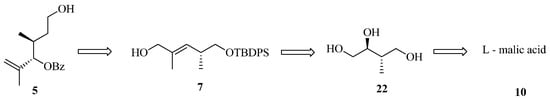
The retrosynthetic strategy for the C9–C13 alcohol fragment 5 is depicted in Scheme 4. The target subunit 5 was envisaged to be derived from allylic alcohol 7, which could, in turn, be prepared from the known triol 22 through a short sequence of transformations starting from L-malic acid (10).
Scheme 4.
Retrosynthetic analysis of the C9–C13 fragment 5 of Pladienolide B (1).
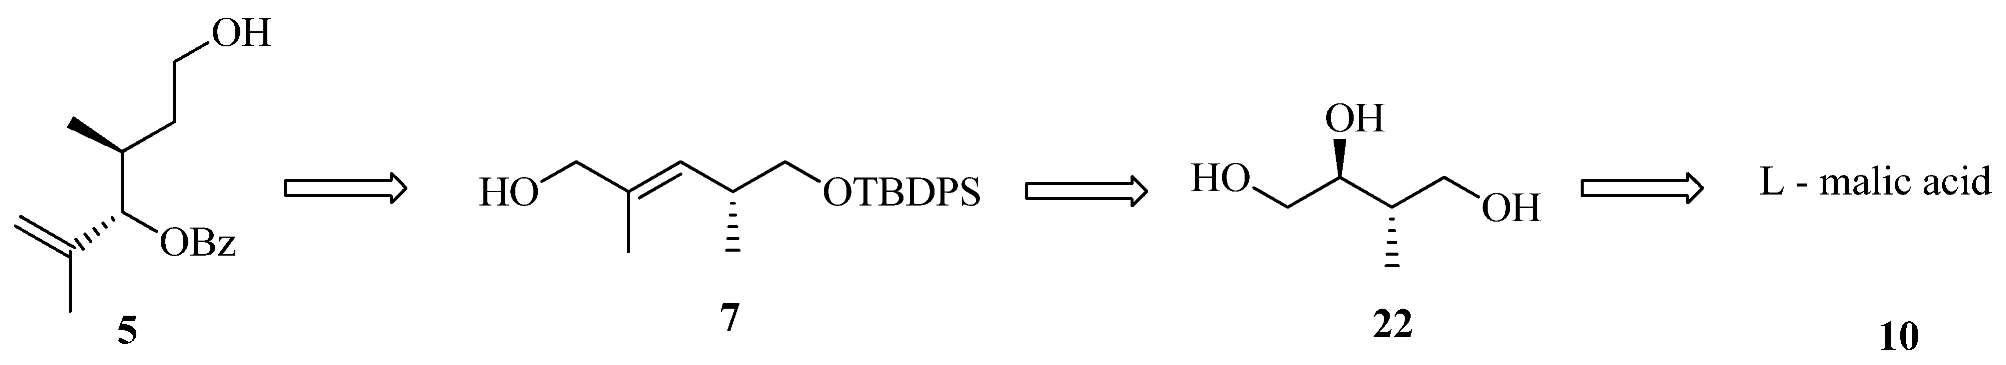
The forward synthesis commenced with the oxidative cleavage of the triol 22 [21] using sodium periodate (NaIO4) in an acetone/water mixture (5:1) to generate the corresponding aldehyde, as shown in Scheme 5. Subsequent Wittig olefination of this intermediate with ethyl 2-(triphenylphosphoranylidene)propanoate yielded the α,β-unsaturated ester 23 in 71% overall yield over two steps. Protection of the secondary hydroxyl group in the compound 23 as a tert-butyldiphenylsilyl (TBDPS) ether using TBDPSCl and imidazole in dichloromethane at 0 °C for 1 h provided the silyl ether 24. Chemoselective reduction of the ester 24 using DIBAL-H in dichloromethane delivered the alcohol 7 in 72% yield. Sharpless asymmetric epoxidation of the allylic alcohol 7 using L-(+)-diethyl tartrate, Ti(OiPr)4, and tert-butyl hydroperoxide (TBHP) in dichloromethane at −20 °C afforded epoxy alcohol 25 with the desired stereochemistry at the epoxide centers. Treatment of the epoxy alcohol 25 under Appel-type conditions afforded the corresponding iodomethyl intermediate 25a. Subsequent zinc-promoted reductive elimination of 25a using zinc powder and NaI in refluxing methanol for 4 h furnished the desired 1,1-disubstituted allylic alcohol 26.
Scheme 5.
Synthesis of C9–C13 fragment 5 of Pladienolide B.
Benzoylation of the allylic alcohol 26 with benzoyl chloride in the presence of triethylamine in dichloromethane at 0 °C for 2 h furnished benzoate ester 27. Finally, fluoride-mediated desilylation of 27 using tetra-n-butylammonium fluoride in THF for 2 h smoothly unmasked the primary hydroxyl group, affording the target C9–C13 alcohol fragment 5.
4. Conclusions
In conclusion, a divergent and stereoselective synthetic strategy was developed for the preparation of the C1–C8 acid fragment 4 and the C9–C13 alcohol fragment 5 of the core moiety 2. Both fragments were accessed from L-Malic acid (10) as a common, inexpensive chiral-pool precursor. The synthesis utilized key asymmetric and stereoselective transformations, including Sharpless asymmetric epoxidation, regioselective epoxide opening, and stereoselective functional-group transformations, to establish the required stereochemical features. This common chiral-pool approach provides a unified route to two stereochemically defined building blocks of Pladienolide B (1) and demonstrates the utility of L-Malic acid (10) for the synthesis of structurally complex natural product fragments.
Supplementary Materials
The following supporting information can be downloaded at https://www.mdpi.com/article/10.3390/org7040049/s1, Copies of 1H NMR and 13 C NMR Spectra of compounds 4–8, 11–13, 15–21, 23–27.
Author Contributions
Conceptualization, M.H.; Methodology, S.K.C., N.A. and M.H.; Formal analysis, S.K.C. and N.A.; Investigation, S.K.C., N.A. and M.H.; Resources, N.A.; Writing—original draft, M.H.; Writing—review & editing, R.R.P.; Supervision, R.R.P.; Project administration, R.R.P.; Funding acquisition, S.K.C. and R.R.P. All authors have read and agreed to the published version of the manuscript.
Funding
This research received no external funding.
Data Availability Statement
The original contributions presented in this study are included in the article/Supplementary Material. Further inquiries can be directed to the corresponding authors.
Acknowledgments
The authors gratefully acknowledge the University Grants Commission (UGC) and the Council of Scientific and Industrial Research (CSIR), New Delhi, for their financial support. The authors also acknowledge Gachon University for its financial support.
Conflicts of Interest
The authors declare no conflicts of interest.
References
- Sakai, T.; Sameshima, T.; Matsufuji, M.; Kawamura, N.; Dobashi, K.; Mizui, Y. Pladienolides, New Substances from Culture of Streptomyces platensis Mer-11107. I. Taxonomy, Fermentation, Isolation and Screening. J. Antibiot. 2004, 57, 173–179. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Sakai, T.; Asai, N.; Okuda, A.; Kawamura, N.; Mizui, Y. Pladienolides, New Substances from Culture of Streptomyces platensis Mer-11107. II. Physico-Chemical Properties and Structure Elucidation. J. Antibiot. 2004, 57, 180–187. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Mizui, Y.; Sakai, T.; Iwata, M.; Uenaka, T.; Okamoto, K.; Shimizu, H.; Yamori, T.; Yoshimatsu, K.; Asada, M. Pladienolides, New Substances from Culture of Streptomyces platensis Mer-11107. III. In Vitro and In Vivo Antitumor Activities. J. Antibiot. 2004, 57, 188–196. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Kotake, Y.; Sagane, K.; Owa, T.; Mimori-Kiyosue, Y.; Shimizu, H.; Uesugi, M.; Ishihama, Y.; Iwata, M.; Mizui, Y. Splicing Factor SF3b as a Target of the Antitumor Natural Product Pladienolide. Nat. Chem. Biol. 2007, 3, 570–575. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Lee, S.C.; Abdel-Wahab, O. Therapeutic Targeting of Splicing in Cancer. Nat. Med. 2016, 22, 976–986. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Jamal-El-Hussein, N.; Chaudhary, S.; Montaudon, E.; Nemati, F.; Genoux, A.; Polveche, H.; El Botti, R.; Barette, C.; Jia, T.; Auboeuf, D.; et al. The SF3B1 Inhibitor Pladienolide B (1) Massively Inhibits DNA Damage Signaling and Repair and Counteracts Resistance to Platinum Salts in Non-Small Cell Lung Cancer. bioRxiv 2026. bioRxiv:2026.02.17.706284. [Google Scholar] [CrossRef] [Scilit]
- Xiao, W.; Yang, L.; Li, Z.; Wang, W.; Liu, Z.; Liu, B.; Qin, J.; Li, Y. Synergistic Effect of Pladienolide B (1) and Cisplatin: Enhancing Autophagy in Hepatoma Cells through the AMPK/mTOR/ULK1 Pathway. Cell Death Discov. 2026, 12, 288. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Guo, Y.; Wang, X.; Du, Y.; Zhao, Y.; Gao, L.; Hao, Y.; Lv, D.; Feng, X.; Zhai, Y.; Zou, B.; et al. The Splicing Factor SF3B1 Confers Ferroptosis Resistance and Promotes Lung Adenocarcinoma Progression via Upregulation of SLC7A11. Cancer Gene Ther. 2024, 31, 1498–1510. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Herrera Lopez, X.I.; Martinez-Perez, K.; Guha Thakurta, S.; Levi, B.P.; Paulo, J.A. Decoding Spliceosome Inhibition: Isobaric Tag-Based Proteomic Profiling of Pladienolide B (1) Treated Human Cell Lines. J. Proteom. 2026, 323, 105565. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Gaunt, M.J.; Jessiman, A.S.; Orsini, P.; Tanner, H.R.; Hook, D.F.; Ley, S.V. Synthesis of the C-1–C-28 ABCD Unit of Spongistatin 1. Org. Lett. 2003, 5, 4819–4822. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Gao, Y.; Hanson, R.M.; Klunder, J.M.; Ko, S.Y.; Masamune, H.; Sharpless, K.B. Catalytic asymmetric epoxidation and kinetic resolution: Modified procedures including in situ derivatization. J. Am. Chem. Soc. 1987, 109, 5765–5780. [Google Scholar] [CrossRef] [Scilit]
- Masamune, S.; Kim, C.U.; Wilson, K.E.; Spessard, G.O.; Georghiou, P.E.; Bates, G.S. Syntheses of macrolide antibiotics I. Methymycin. J. Am. Chem. Soc. 1975, 97, 3512–3513. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Kanada, R.M.; Itoh, D.; Nagai, M.; Niijima, J.; Asai, N.; Mizui, Y.; Abe, S.; Kotake, Y. Total synthesis of the potent antitumor macrolides pladienolide B and D. Angew. Chem. 2007, 119, 4428–4433. [Google Scholar] [CrossRef] [Scilit]
- Ghosh, A.K.; Anderson, D.D. Enantioselective Total Synthesis of Pladienolide B: A Potent Spliceosome Inhibitor. Org. Lett. 2012, 14, 4730–4733. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Kumar, V.P.; Chandrasekhar, S. Enantioselective Synthesis of Pladienolide B and Truncated Analogues as New Anticancer Agents. Org. Lett. 2013, 15, 3610–3613. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Arai, K.; Buonamici, S.; Chan, B.; Corson, L.; Endo, A.; Gerard, B.; Hao, M.-H.; Karr, C.; Kira, K.; Lee, L.; et al. Total Synthesis of 6-Deoxypladienolide D and Assessment of Splicing Inhibitory Activity in a Mutant SF3B1 Cancer Cell Line. Org. Lett. 2014, 16, 5560–5563. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Sim, J.; Jang, E.; Kim, H.J.; Jeon, H. Total Syntheses of Pladienolide-Derived Spliceosome Modulators. Molecules 2021, 26, 5938. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Selka, A.; Abidli, A.; Schiavo, L.; Jeanmart, L.; Hanquet, G.; Lubell, W.D. Recent Advances in Sustainable Total Synthesis and Chiral Pool Strategies with Emphasis on (−)-Sclareol in Natural Products Synthesis. Eur. J. Org. Chem. 2025, 28, e202400983. [Google Scholar] [CrossRef] [Scilit]
- Yoo, M.; Krische, M.J. Total Synthesis of the Spliceosome Modulator Pladienolide B (1) via Asymmetric Alcohol-Mediated syn- and anti-Diastereoselective Carbonyl Crotylation. Angew. Chem. Int. Ed. 2021, 60, 13923–13928. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Rhoades, D.; Rheingold, A.L.; O’Malley, B.W.; Wang, J. Expedient Total Syntheses of Pladienolide-Derived Spliceosome Modulators. J. Am. Chem. Soc. 2021, 143, 4915–4920. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Yadav, J.S.; Sengupta, S. The Formal Total Synthesis of FR252921–An Immunosuppressant. Eur. J. Org. Chem. 2013, 376–388. [Google Scholar] [CrossRef] [Scilit]
Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2026 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license.