Article

Native Tapioca Starch Agglomerated Using Its Gelatinized Dispersion for Tablet Production by Direct Compression


by
Rapee Jarungsirawat
1,
Chaipat Siriwachirachai
1 and
Thaned Pongjanyakul
1,2,*
1
Faculty of Pharmaceutical Sciences, Khon Kaen University, Khon Kaen 40002, Thailand
2
Center for Research and Development of Herbal Health Products, Khon Kaen University, Khon Kaen 40002, Thailand
*
Author to whom correspondence should be addressed.
Sci. Pharm. 2026, 94(3), 73; https://doi.org/10.3390/scipharm94030073
Submission received: 24 July 2026 / Revised: 21 August 2026 / Accepted: 26 August 2026 / Published: 31 August 2026

Abstract

This study aimed to investigate the physical properties of native tapioca starch (TS) ag-glomerated using its gelatinized dispersion as an agglomerating agent and to evaluate the performance of the resulting agglomerate tablets. The results demonstrate that TS agglomerated with gelatinized tapioca starch (GTS) exhibited increased particle strength as GTS content increased. Moreover, particle flowability was enhanced compared with that of native TS. The compressibility of TS agglomerates increased with higher GTS content, thereby reducing resistance to volume reduction and enhancing the plasticity of particle deformation under pressure, resulting in the greater tensile strength of the tablets. GTS showed superior performance to polyvinylpyrrolidone and sodium alginate at the concentration of 2% w/w in terms of Carr’s index and tablet hardness. Propranolol HCl (PNL) tablets prepared from agglomerates with GTS exhibited higher hardness than those without GTS, with tablet hardness increasing proportionally to GTS content. Additionally, GTS facilitated faster tablet disintegration, thereby accelerating PNL dissolution. In drug-loading tests, agglomerate tablets containing 3% GTS maintained acceptable physical properties when the PNL content did not exceed 20% w/w. Higher PNL loading may be achievable by increasing compression pressure. These findings indicate that native TS agglomerated with GTS is a promising tablet diluent for direct compression.

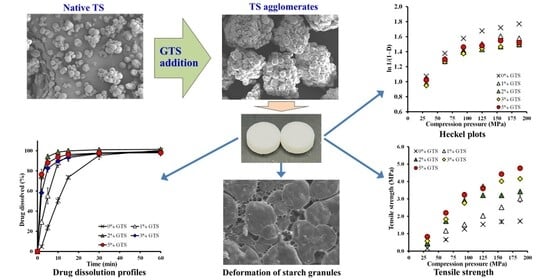
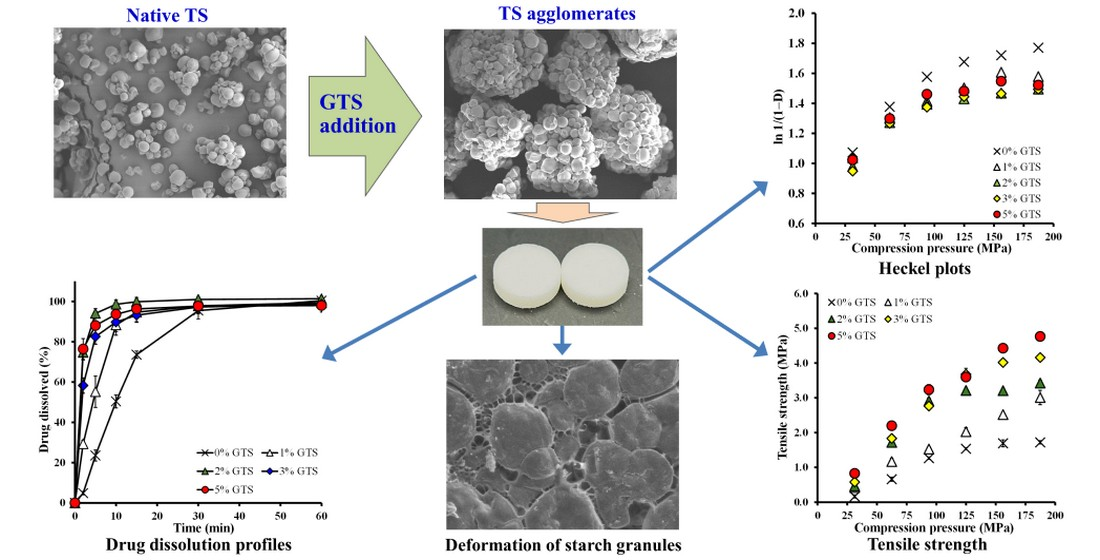
Graphical Abstract

1. Introduction

Tapioca starch (TS), derived from the cassava (Manihot esculenta (L.) Crantz) root, is an abundant material produced in northeast Thailand [1]. Starches consist of two main components: amylose and amylopectin. Amylose is a linear chain of glucose molecules linked by α-1-4 glycosidic bonds, while amylopectin is a branched chain featuring α-1-6 glycosidic linkages [2,3,4]. Native TS can serve as a diluent, binder or disintegrant in tablet dosage forms [1,3,5]. It has low water solubility and does not swell at ambient temperature [6]. When heated above the gelatinization temperature, however, it can be hydrated and swollen to form a clear dispersion [7,8]. This gelatinized TS dispersion has been used as a binder in the wet granulation method for tablet preparation [9]. However, using native TS as the primary diluent for tablet preparation by direct compression is limited due to its insufficient flowability and compressibility [8,10,11,12].
Various chemical and physical modifications have been investigated to improve the physicochemical properties of starch for pharmaceutical uses [2,13,14,15]. Acid modification followed by spray drying increases the gelatinization temperature and relative crystallinity of TS, resulting in enhanced compressibility compared to the native form [11]. However, using high-temperature spray drying may cause gelatinization during the drying process. As a result, tray drying and particle size selection were also employed in the acid modification process. Despite these approaches, the modified TS exhibited poor flowability due to its bulky, cohesive nature. However, the hardness of tablets made from this modified TS was higher than that of the native TS [8]. Therefore, an agglomeration method was developed to enlarge the particles of acid-modified TS to enhance flowability [12].
Agglomeration increases the size of small particles by using an agglomerating agent that coats them, enhancing adhesion. Particle adhesion is mediated by mechanisms such as liquid bridges, solid bridges, and interlocking bonds [16]. The equipment used for agglomeration closely resembles that used in wet granulation for tablet production. Previous research has demonstrated the successful formation of particle agglomerates from drug powders and polymeric composites using polyvinylpyrrolidone (PVP) as the agglomerating agent [12,17,18,19]. Acid-modified TS agglomerated with PVP exhibits improved flowability and compressibility. Furthermore, PVP has been used to enhance the flowability of native TS, with the compressibility of the resulting agglomerates depending on PVP concentration [12]. These findings indicate that native TS can be agglomerated without prior modification, thereby avoiding the complexity and time requirements of starch alteration.
Native TS can also be agglomerated using sodium alginate (SA), which improves particle flowability. The hardness of agglomerate tablets increases with higher compression pressure, and this effect is influenced by both SA concentration and viscosity grade [20]. This indicates that the properties of TS agglomerates are affected by the characteristics of the agglomerating agent used. Thus, it is worthwhile to explore readily available and inexpensive materials for agglomeration. Using a gelatinized dispersion of TS for this purpose is a logical choice, as it can act as a binder in wet granulation [5,9], providing a thin film on the starch granules’ surfaces to facilitate agglomeration. Moreover, tablets made from gelatinized TS (GTS) powder exhibit greater hardness than those made from native TS [21]. This property may further enhance the compressibility of TS agglomerates prepared with GTS. Moreover, the use of GTS may reduce manufacturing costs and avoid the use of synthetic materials in the production of TS agglomerates.
This study aimed to prepare and evaluate TS agglomerates using GTS dispersion as the agglomerating agent. TS agglomerates with varying GTS contents were produced, and their characteristics, including particle morphology, particle strength, flowability, and compressibility, were assessed. Furthermore, the effect of drug loadings, also known as the carrying capacity or dilution potential of the diluents, on the physical properties and drug dissolution profiles of tablets prepared using TS agglomerates was investigated.

2. Materials and Methods

2.1. Materials

Native TS was purchased from Kaensiri Starch Co., Ltd. (Khon Kaen, Thailand). Propranolol hydrochloride (PNL) and magnesium stearate were obtained from Changzhou Yabang Pharmaceutical Co., Ltd. (Changzhou, China) and Mallinckrodt Inc. (St. Louis, MO, USA). Colloidal silicon dioxide (Aerosil 200) was received from Maxway Co., Ltd. (Bangkok, Thailand). Magnesium stearate and colloidal silicon dioxide were passed through a 180-μm sieve before use. All other reagents were of analytical grade and used as received.

2.2. Preparation of TS Agglomerates

Native TS in 0, 2, 4, 6, or 10 g was dispersed in 50 mL of purified water. The starch suspension was incubated at 85 °C in a water bath for 10 min to produce a GTS dispersion. Once the GTS dispersion became transparent, it was cooled to room temperature before use. Native TS weighing 200, 198, 196, 194, or 190 g was crushed using a mortar and pestle. The GTS dispersion was then combined with the crushed native TS in the mortar to create mixtures containing 0, 1, 2, 3, or 5% w/w GTS, respectively. Additional water may be needed to achieve an appropriate damping mass. The mixing process lasted 5 min to ensure even distribution of the GTS throughout the damping mass. After mixing, the damping mass was passed through a 1.7 mm sieve and dried at 55 °C in a hot air oven. The dried particles obtained were gently ground using a mortar and pestle. Small particles were passed through a 150-μm sieve using a spatula to select the desired particle size. TS agglomerates with a particle size range of 75–150 μm were collected using a sieve shaker (Retsch GmbH AS200 control, Haan, Germany), and the percentage yield of the obtained agglomerates was also determined. The agglomerates with GTS showed a tacky property, as observed visually, especially with 5% GTS. To solve this problem, colloidal silicon dioxide, which was passed through a 180-μm sieve, was added at a very low concentration (0.1% w/w) to all agglomerates using GTS to adsorb residual water and prevent tackiness. Afterward, the agglomerates were stored at room temperature (26–28 °C) in a desiccator containing silica gel beads before use.
The application of colloidal silicon dioxide was evaluated in this study. Colloidal silicon dioxide is commonly employed as a glidant in tablet formulations. The influence of this excipient at a concentration of 0.1% w/w on the flowability and tablet hardness of agglomerates was preliminarily assessed, despite its low content. Results indicated that the addition of 0.1% colloidal silicon dioxide produced only minor changes in flowability and tablet hardness compared to agglomerates without this excipient. Consequently, 0.1% colloidal silicon dioxide was incorporated into the agglomerates. However, subsequent analyses, including scanning electron microscopy (SEM), particle strength, compressibility assessments using Heckel and Kawakita models, and the effect of drug loadings, were performed on agglomerates without the addition of 0.1% colloidal silicon dioxide.

2.3. Characterization of TS Agglomerates

2.3.1. Particle Morphology Studies

The particle morphology of the native TS and its agglomerates without 0.1% colloidal silicon dioxide was examined using scanning electron microscopy (SEM). The samples were mounted onto a dummy substrate using adhesive carbon tape. They were then coated with gold in a vacuum system and observed under a Hitachi S-3000N scanning electron microscope in Tokyo, Japan.

2.3.2. Measurement of Particle Strength

The strength of the agglomerates was measured using a previously reported method [17,22]. The larger particle size of the agglomerates without 0.1% colloidal silicon dioxide was utilized. The damping mass of TS mixed with GTS was passed through a 1.7 mm sieve and dried at 55 °C in a hot-air oven. A texture analyzer (TA.XT Plus, Stable Micro Systems, Godalming, UK) equipped with a 50 kg load cell and a 6 mm-diameter cylindrical probe was used for the measurements. The agglomerate was placed on the platform at room temperature, and the probe was positioned to contact the agglomerate’s surface before moving downward at a constant rate of 1.0 mm/s. The probe was automatically withdrawn once the agglomerates were compressed to 50% of their original height. The force and percent displacement were plotted, with the maximum force at 50% displacement representing the strength of the agglomerates being reported.

2.3.3. Determination of Densities and Flowability

The agglomerated starches (10 g) were weighed and gently poured into a 50 mL cylinder. The sample volume was measured, and the weight divided by volume was computed as the bulk density (Db). The cylinder containing the sample was then tapped with a 1.5-inch height until the sample volume was constant. The final volume was used to calculate the tapped density (Dt). The true density of the sample was determined using a liquid-displacement method with a 5 mL pycnometer, using acetone as the liquid to prevent the agglomerate’s ingredients from dissolving. A weighed pycnometer was filled with 1 g of the samples and weighed accurately. The pycnometer with the samples was filled using acetone to the maximum volume, and the total weight of the filled pycnometer was determined. The weight of the acetone in the pycnometer without the sample was also measured. The true density was calculated using the equation as follows: true density = (WA × WS)/(5 (WA − WSA + WS)) [23], where WA, WS, and WSA represent the weights of acetone, samples, and sample-acetone mixture in the pycnometer, respectively.
The flowability of the agglomerates was assessed in terms of the angle of repose and Carr’s index. The angle of repose was measured by continuously pouring the agglomerates through a funnel onto a smooth surface, forming a conical pile of fixed height. The height (h) and radius (r) of the pile were recorded, and the angle of repose (θ) was calculated using the equation: θ = arctan (h/r) [8,24]. Carr’s index was related to the bulk and tapped densities, and could be calculated using the formula: Carr’s index (%) = 100 × (Dt − Db)/Dt [8,24].

2.3.4. Compressibility of TS Agglomerates

The agglomerates (400 mg) without 0.1% colloidal silicon dioxide were weighed and filled into a 10 mm-diameter die. Then the agglomerates were compressed into tablets using a 10 mm-diameter flat-faced punch at various compression pressures (31.2–249.8 MPa) in a hydraulic press (Retsch PP 25 Pellet Press, Haan, Germany). The die and punches had been lubricated using magnesium stearate powder before use. Three tablets were compressed for each compression pressure. The out-die thickness of each tablet was measured immediately after ejection using a digital Vernier caliper with an accuracy of ±0.01 mm. The tablet density and volume under compression pressure were calculated. Moreover, tensile strength of the tablets obtained was also computed using the equation: tensile strength (σt) = 2F/πDT [25,26], where F is the break force of flat faced tablets subject to diametrical compression (referring to a tablet hardness), π is a constant of 3.143, D is a tablet diameter, and T is tablet thickness.
The compressibility of the agglomerates was studied using the Heckel and Kawakita plots. The Heckel analysis describes the compression characteristic of a powder in terms of its relative density under applied pressure. The densification of the bulk powder under pressure follows first-order kinetics that can be expressed by the equation [27]:
ln(1/(1 − D)) = KP + A
where D represents the relative density of the tablet (the ratio of tablet density to the true density of the agglomerates) under compression pressure (P). The relationship between ln(1/(1 − D)) and P was plotted, and the slope and intercept were determined [27,28,29]. K is the slope of the linear portion of the Heckel plot, and the reciprocal of K provides the mean yield pressure (Py). The A value describes the densification of the samples due to initial particle rearrangement (Da), which can be calculated using an additional equation:
Da = 1 − e−A
The Kawakita analysis is used to investigate particle deformation under compression pressure by examining a powder volume change at various compression pressures. The equation was presented as follows [28,29]:
P/C = (P/a) + (1/ab)
C = (V0 − V)/V0
where P is the compression pressure, and C describes the volume decrease after applying compression pressure. V0 is the initial volume of the powder filled into the die, which was computed from the bulk density. V is the tablet volume after compression, determined from tablet thickness. The a value shows the initial porosity, which decreases when pressure is applied. The b value is proposed to be inversely related to the yield strength of particles (Pk). Data for P/C and P were plotted, and the slope (1/a) and intercept (1/ab) constants were obtained.

2.4. Preparations of Agglomerate Tablets

A tablet was compressed at low compression pressure (12.3 MPa) using agglomerates with various GTS as a diluent. Each tablet contained 40 mg of PNL along with 210 mg of TS agglomerates. This mixture was combined in a roto-mixer for 3 min. Then, 1% w/w magnesium stearate was added, and the mixture was mixed continuously for an additional 2 min. A total of 252.5 mg of this mixture was filled into a 10 mm diameter flat-faced die, compressed at 12.3 MPa using a hydraulic press (Model 3126, Shimadzu, Kyoto, Japan) without holding time, and the PNL tablets obtained were stored in a desiccator prior to testing. Moreover, tablets without PNL were also prepared for comparison of hardness with those containing PNL.
The effect of PNL loadings on the characteristics of the agglomerate tablets was investigated using the TS agglomerates with 3% GTS, and scaled-up preparation for a 1 kg batch was performed. The native TS 30 g was dispersed in 250 mL of purified water, and the starch suspension was incubated at 85 °C in a water bath for 10 min to produce a GTS dispersion. The native TS (970 g) and the GTS dispersion were blended in a mixer machine (Kenwood Major, Birmingham, UK) to achieve a wet mass. The damping mass was passed through a 1.7 mm sieve using an oscillating granulator (Erweka GmbH AR400, Heusenstamm, Germany) and then dried at 55 °C in a hot-air oven. The dried particles were then sieved using a 150-μm sieve, and the agglomerates (particle size 75–150 μm) were collected using a sieve shaker (Retsch GmbH AS200 control, Haan, Germany). The moisture content of the agglomerates was measured using an Aczet MB50 Moisture Analyzer (Mumbai, India). The agglomerates (2 g) were placed in a sample pan and heated to 90 °C until a constant weight was reached. The moisture content (% w/w) was computed from the weight loss during heating.
The model drug used in this study was PNL. Each tablet weighed 350 mg and contained varying amounts of PNL (0, 10, 20, 30, and 40% w/w) and TS agglomerates. Both components were mixed in a roto-mixer for 10 min. Following this, 1% w/w magnesium stearate was incorporated into the mixture, which was remixed for another 5 min. The tablets were then compressed using a single-punch tableting machine (Yeo Heng Co., Ltd., Bangkok, Thailand) with a 10 mm flat-faced die and accompanying punches, producing a batch of 500 tablets at a compression rate of 45 tablets/min. This study implemented control over tablet thickness and weight. The tablets were stored in a desiccator with silica gel beads for further examination. The physical properties and drug dissolution of the tablets were assessed.

2.5. Evaluations of Tablets

2.5.1. Thickness, Hardness, and Disintegration Time

The thickness and hardness of the tablets were evaluated using a Vernier caliper and a tablet hardness tester (Model 40-2100 VK200 VanKel®, Cary, NC, USA). The disintegration time of the tablets was measured using a basket rack assembly disintegration test apparatus (Model ZT-324, Erweka America Inc., Edison, NJ, USA). The media used were 0.1 N HCl and a pH 6.8 phosphate buffer, maintained at 37.0 ± 0.5 °C. Each tablet was placed in the basket without a dish, and the disintegration time was recorded when the tablet was entirely liberated from the basket screen.

2.5.2. Weight Variation, Friability, and PNL Content

For the PNL loading study, weight variation, percent friability, and PNL content were also determined. Twenty tablets were weighed using an analytical balance, and the results were reported as the average weight with standard deviation (SD) and relative standard deviation (RSD). A friability test was also performed. Excess powder was carefully removed from the tablet surfaces prior to testing. At least 6.5 g of tablets were weighed and placed in the drum of a tablet friability instrument (Model 45-2200 VanKel®, Cary, NC, USA). The drum containing the tablets was rotated for 100 revolutions over 4 min. Afterward, the tablets were removed, and excess dust was cleared away. The remaining weight of the tablets was accurately measured, and the percent friability (weight loss) was calculated. Additionally, the PNL content of the tablets was determined by soaking the tablets in 100 mL of 0.1 N HCl for 1 h. The solution was then filtered through a 0.45 µm cellulose acetate membrane. The clear filtrate was analyzed for PNL concentration using a UV–visible spectrophotometer (Shimadzu UV1201, Kyoto, Japan) at 289 nm.

2.5.3. PNL Dissolution

The drug dissolution study of the tablets was conducted with a USP dissolution apparatus I (basket method) (Hanson Research 72RL, Chatsworth, CA, USA). The dissolution medium (750 mL) was 0.1 N HCl, maintained at 37.0 ± 1.0 °C. The basket rotated at 50 revolutions per minute. Samples were collected at designated intervals, and each sample was replaced with an equal volume of fresh medium. The PNL concentrations in the samples were analyzed using a UV–visible spectrophotometer (Shimadzu UV1201, Kyoto, Japan) at 289 nm. The time to achieve 50% dissolution of the drug content (T50%) was calculated to compare the tablets’ dissolution profiles.

2.6. Statistical Analysis

The statistical differences in particle strength and tablet properties, including hardness, disintegration time, and T50% values, were analyzed using one-way analysis of variance (ANOVA) with Tukey’s honest significant difference test for multiple comparisons and Student’s t-test for two-sample comparisons. All statistical analyses were performed using IBM SPSS Statistics for Windows, Version 19.0 (IBM Corp., Armonk, NY, USA), with significance set at p < 0.05.

3. Results and Discussion

3.1. Particle Morphology and Strength

The TS agglomerates created using GTS were successful in this study, with the selected agglomerate sizes ranging from 75 to 150 μm. In the preparation process in Section 2.2, GTS content ranging from 1 to 3% yielded a suitable viscosity for the GTS dispersion for TS agglomeration. However, the higher viscosity of the 5% GTS used made sieving difficult in both wet and dry conditions, but this formulation could still produce TSS agglomerates. Thus, GTS content higher than 5% was not recommended for this purpose.
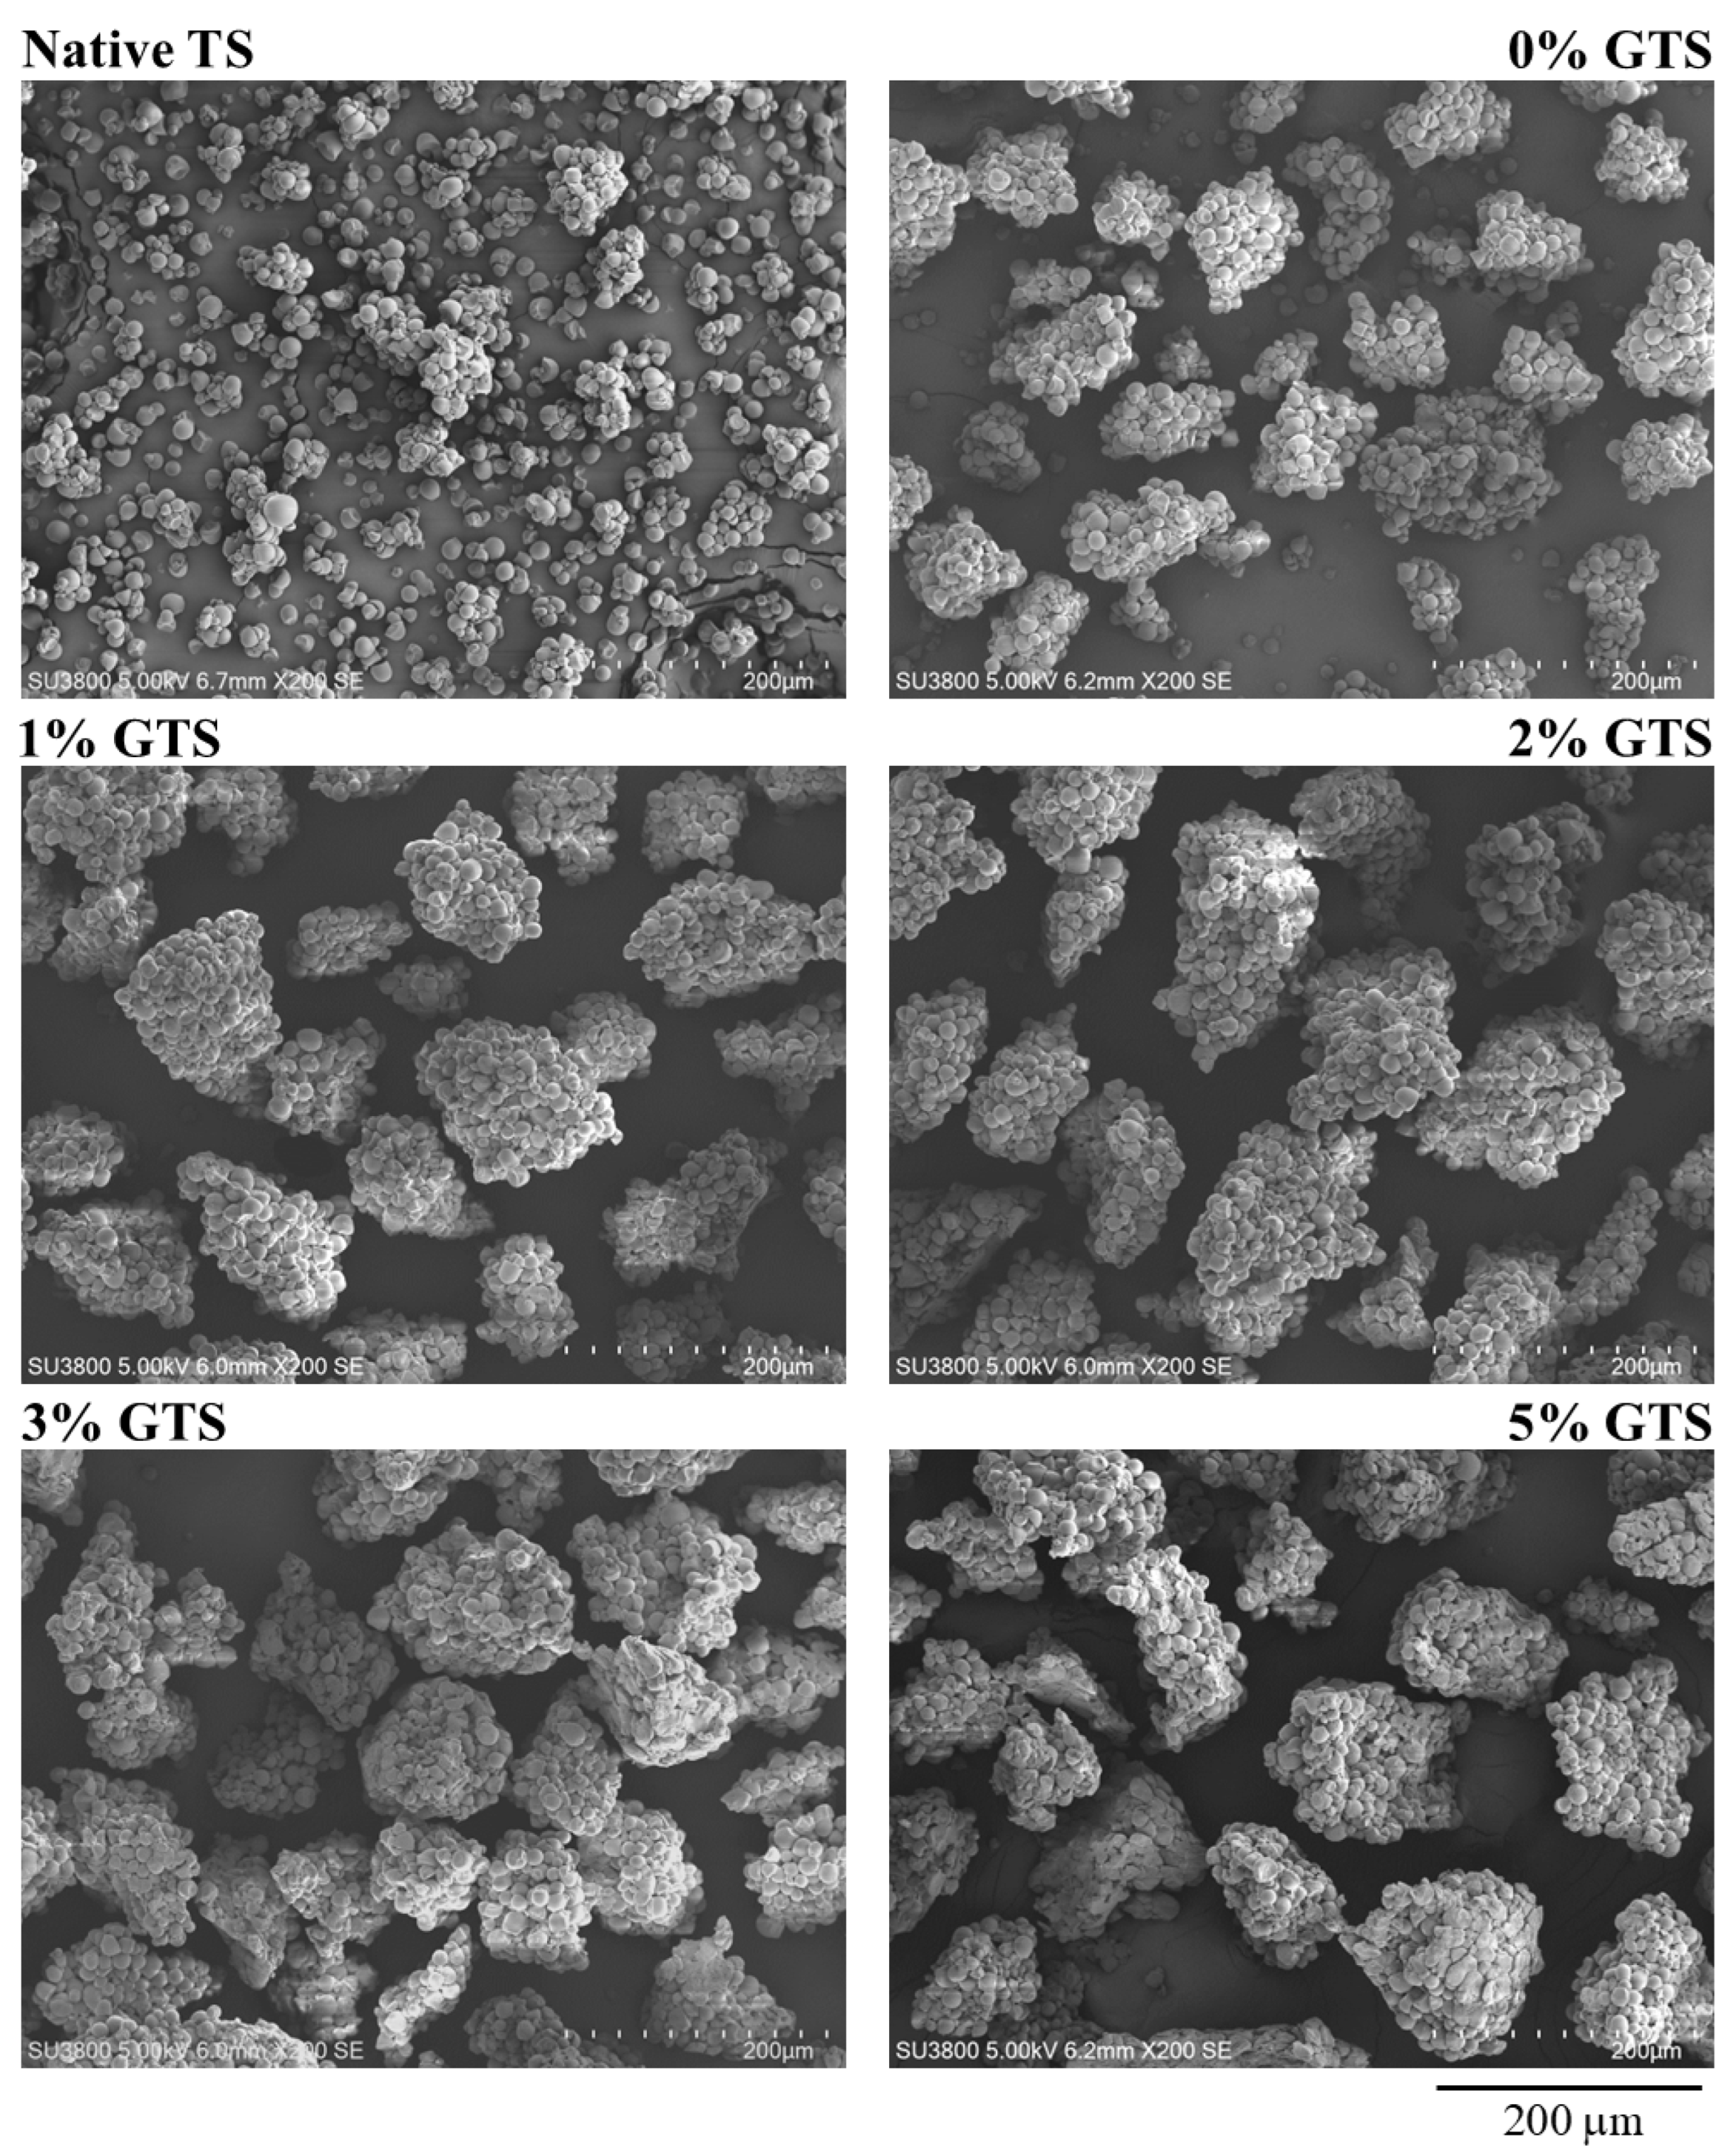
The TS agglomerates created using GTS were successful in this study, with the selected agglomerate sizes ranging from 75 to 150 μm. Figure 1 shows the particle morphologies of both the native TS and the agglomerates, as observed by scanning electron microscopy (SEM). The native TS exhibited small granules along with some aggregates of starch granules. Using purified water (0% GTS) facilitated the agglomeration of TS. Concentrations of GTS ranging from 1% to 5% acted as effective agglomerating agents, producing agglomerates with similar morphologies.
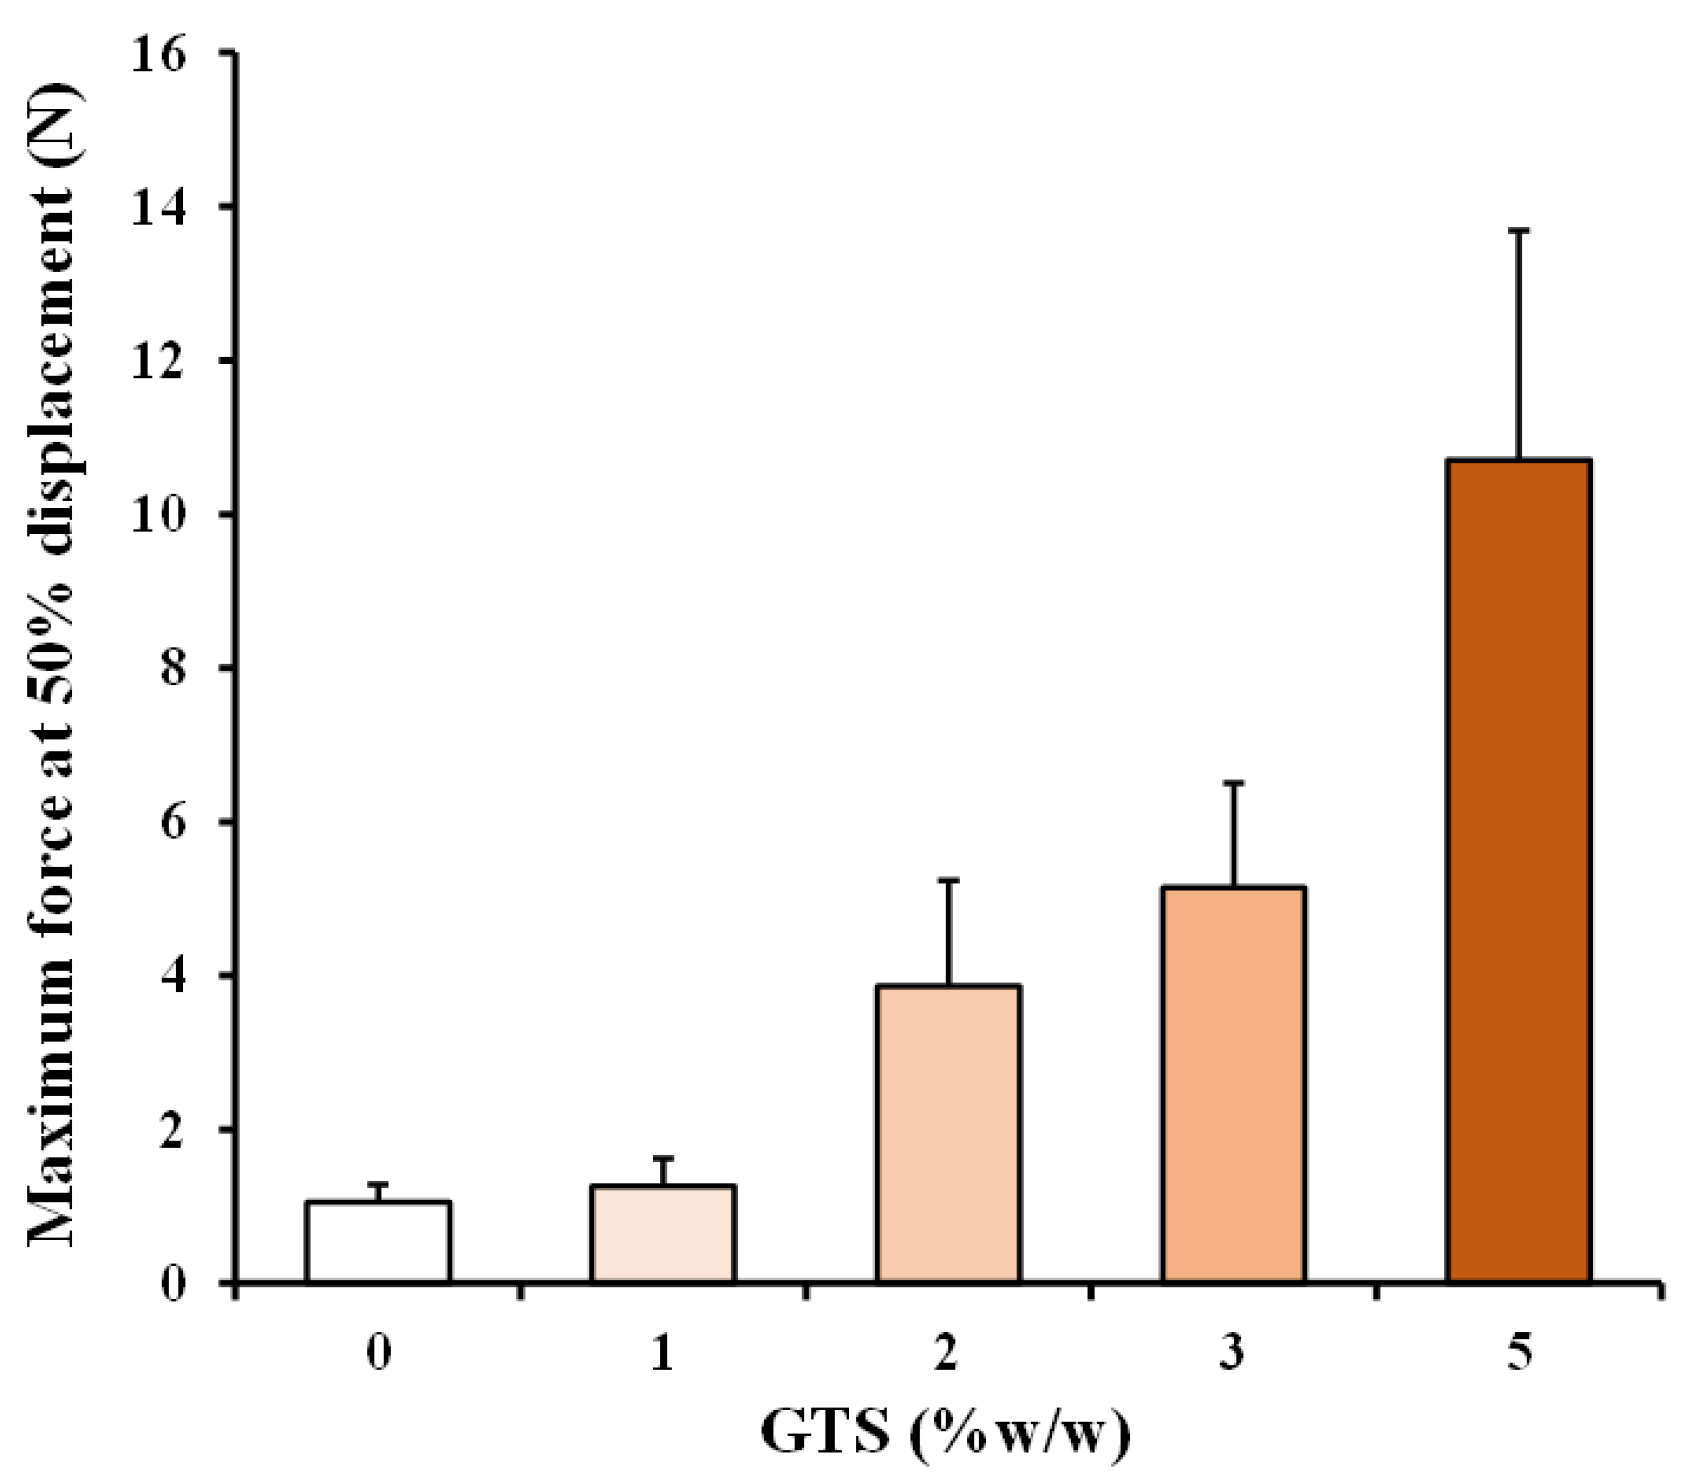
The yields of the selected agglomerates are detailed in Table 1. An increase in GTS content led to a higher agglomerate yield. This finding aligns with previous reports indicating that the yield of larger agglomerates increases with a higher concentration of agglomerating agents [22]. Figure 2 illustrates the strength of the agglomerates, as measured by the maximum force at 50% displacement. The TS agglomerates prepared with purified water showed the weakest strength. As the GTS concentration increased, the strength of the TS agglomerates also increased. Notably, the agglomerates with 5% GTS demonstrated a significantly higher maximum force (p < 0.05) than those with lower GTS concentrations. Therefore, the agglomerates containing 5% GTS were found to be the most robust in this study.
In the TS agglomeration process, the important step is the accumulation of GTS as a thin film on the surface of starch granules. This step reduced the interparticle distance, increased the contact area, and promoted adhesion of starch granules, thereby generating capillary forces and strong solid bridges due to binder hardening after drying [16,30]. Thus, the GTS films around the starch granules acted as an agglomerating agent, enlarging the agglomerates. The increase in GTS content could strengthen the solid bridges between starch granules, leading to stronger agglomerates. Moreover, the stronger agglomerates obtained with higher GTS content led to a higher % yield of particles in the 75–150 micrometer size range, due to less cracking during sieving for size selection.

3.2. Densities and Flowability of the Agglomerates

The bulk and tapped densities of the agglomerates are presented in Table 1. Agglomerates made with purified water exhibited the highest densities. Similar densities were obtained from the TS agglomerates with GTS, with the bulk density being 0.39–0.40 g cm−3, and the tapped density (0.44–0.46 g cm−3) was higher than the bulk density. The true density of all agglomerates, measured using a liquid-displacement method, ranged from 1.516 to 1.548 g cm−3, as shown in Table 1. The bulk and tapped densities could be used to compute Carr’s index (Table 1). The agglomerates formed by water showed the highest Carr’s index value. In contrast, the agglomerates using GTSs had a lower Carr’s index, ranging from 12.1% to 13.5%, which was not influenced by GTS content (Table 1). All agglomerates showed good flowability [31] as evaluated by Carr’s index. The angle of repose for the agglomerates made with purified water was 36.8 degrees, while those with GTSs ranged from 37.8 to 41.3 degrees. These angles indicate a moderate level of particle flowability [32]. However, both parameters of the agglomerates were better than those of the native TS, which was reported to have a Carr’s index of 24.2% and a repose angle of 47.7° in a previous study [8]. Carr’s index reflects the cohesiveness and friction of powders under static conditions [33], while the angle of repose measures the dynamic friction properties of the particles [32]. Therefore, this finding suggests that using the gelatinized form of native TS for agglomeration can reduce the cohesiveness and interparticulate friction forces of starch granules, ultimately enhancing particle flowability. Nevertheless, it could be observed that increasing GTS content did not affect the Carr’s index and repose angle values in this study because the flow properties of powders depend upon the particle size [34]; the use of the same size range of the agglomerates may provide similar flowability characteristics.

3.3. Compressibility of the Agglomerates

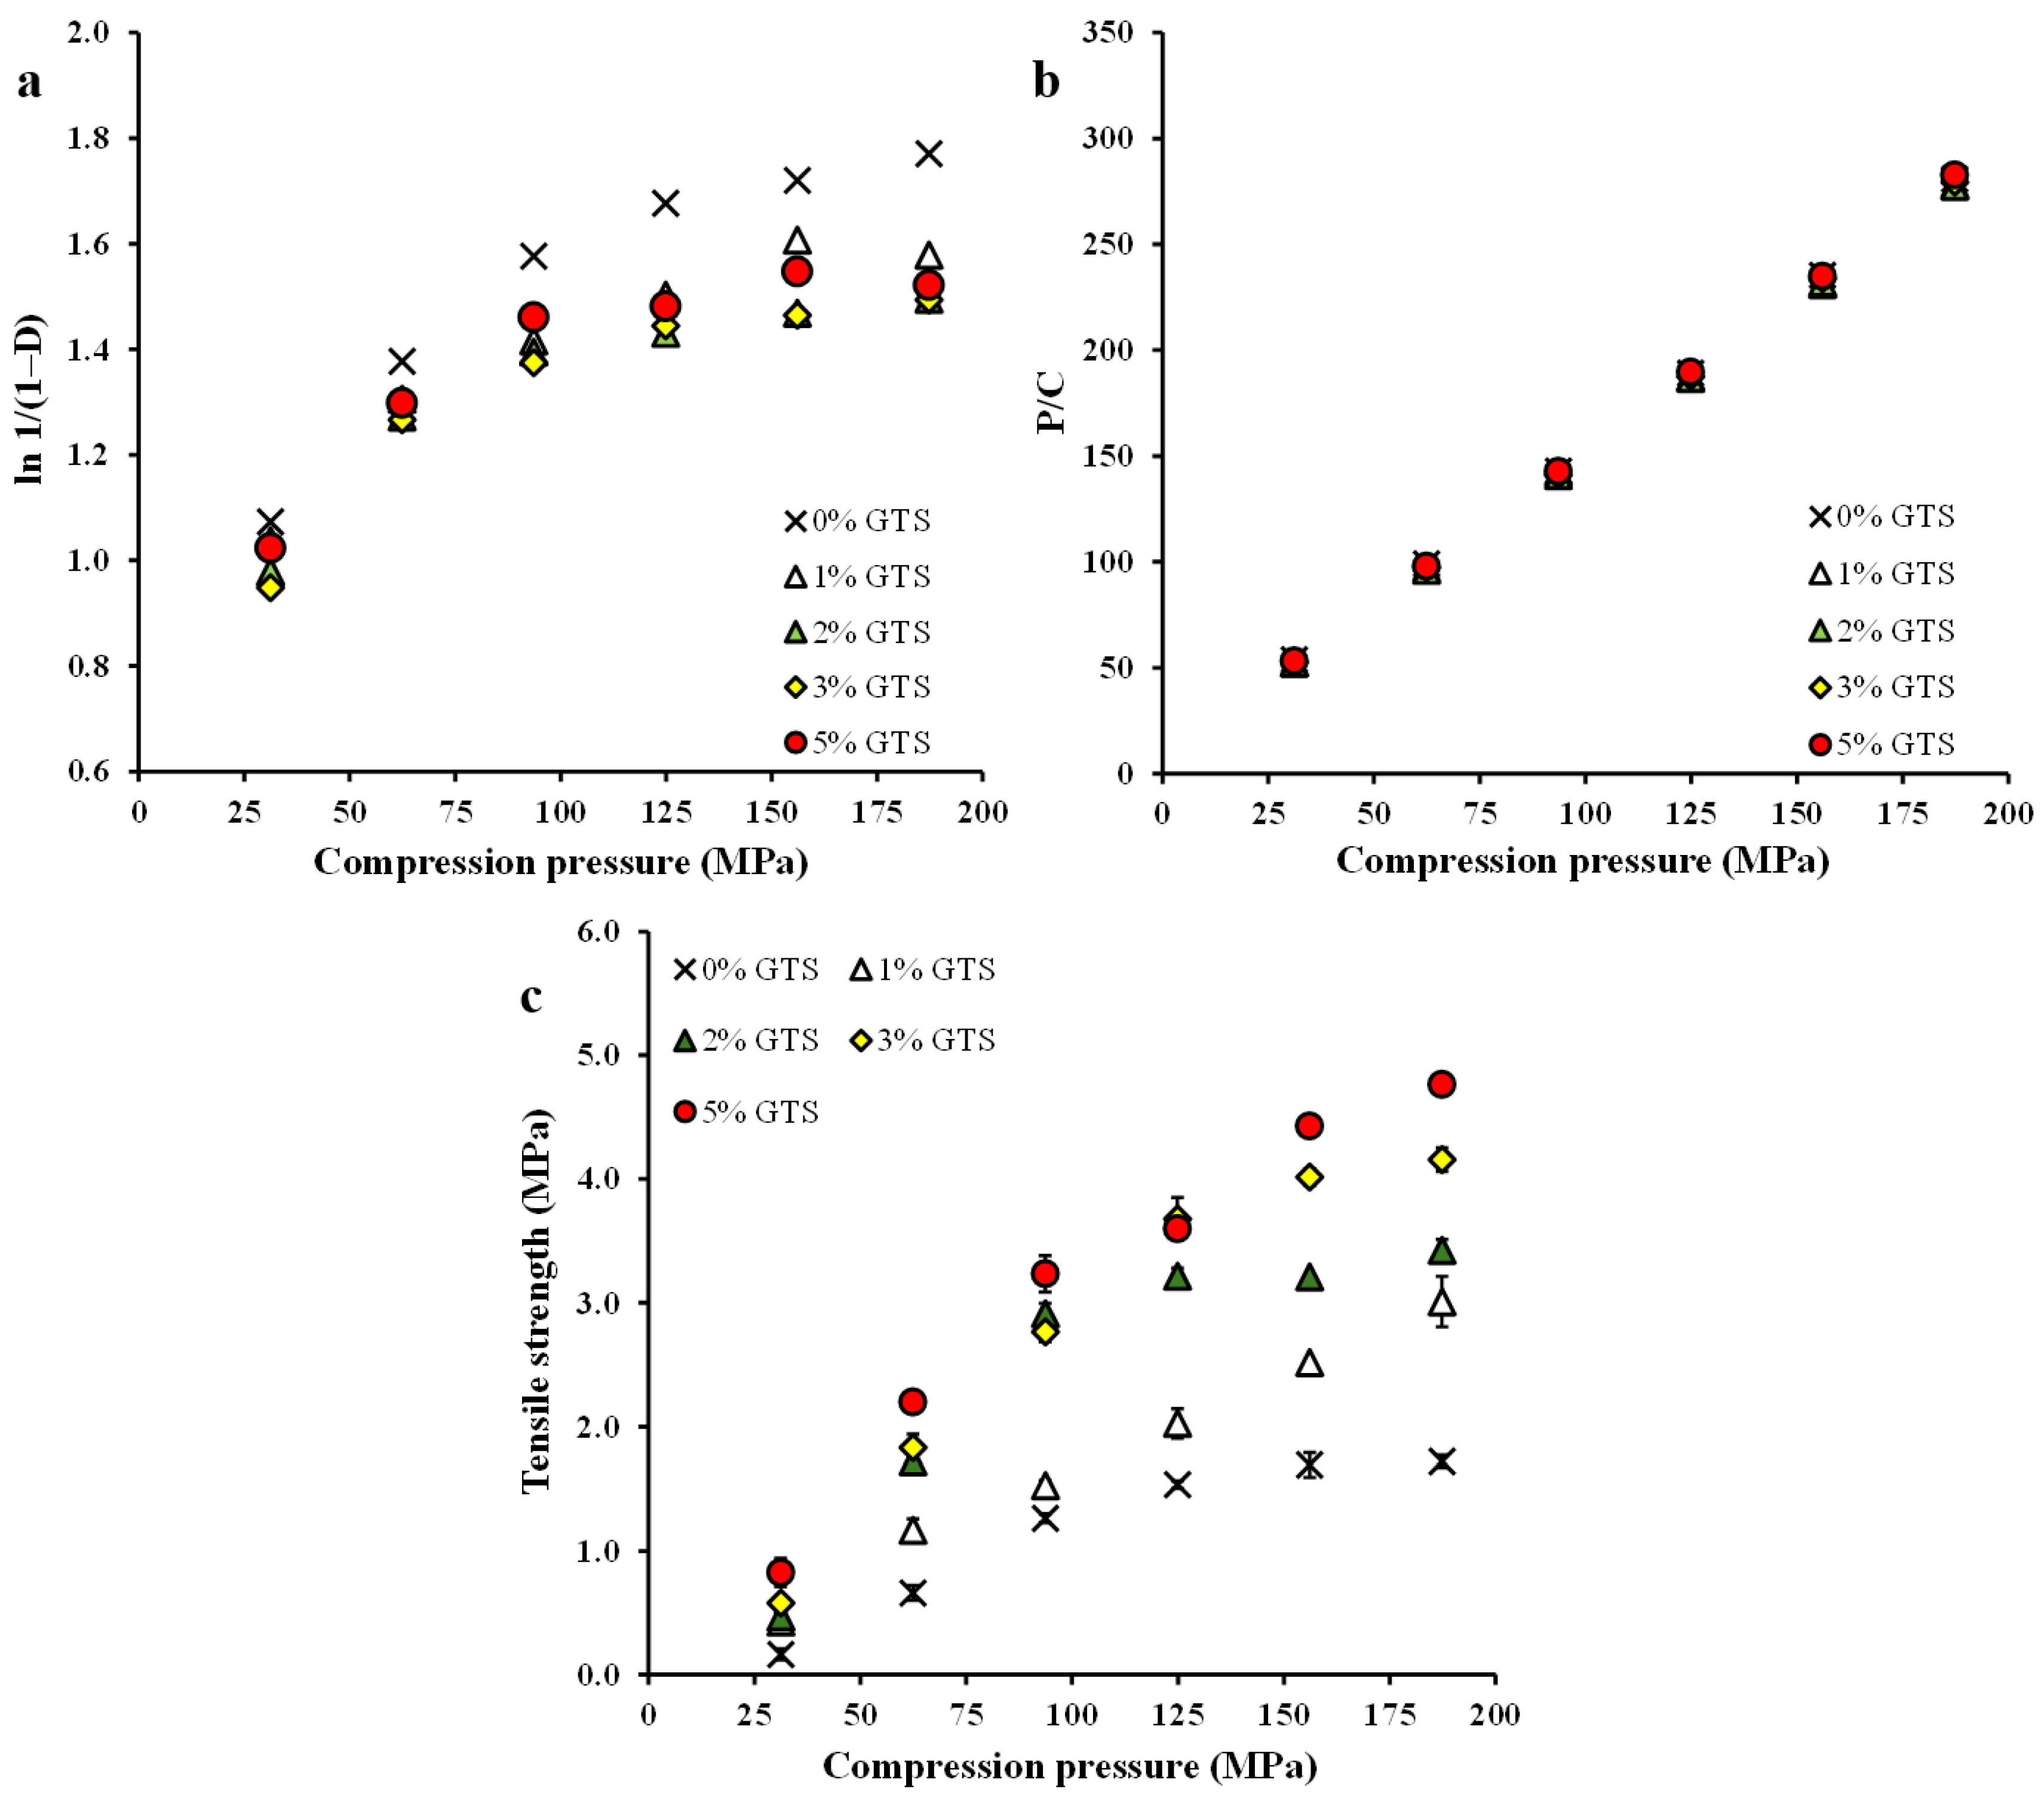
The Heckel and Kawakita plots were used to evaluate the particle deformation under compression pressures. For the Heckel plots, the relationship between ln 1/(1 − D) and compression pressures of the agglomerates using 0–5% GTS is presented in Figure 3a. This relationship showed a good fit with R2 higher than 0.93 in the pressure range of 31.2–93.7 MPa. The parameters involved in this plot are listed in Table 2. The K value of the agglomerates without GTS was 0.643 × 10⁻² 1/MPa, and the Py (yield pressure) was found to be 155.52 MPa. The addition of 1% GTS decreased the K value but increased the Py value compared with those without GTS. The greater the K value, the higher the plastic deformation, and the lower the yield pressure. The K value increased with increasing GTS added, leading to a decrease in Py value. This result suggested that TS agglomeration with GTS led to particle deformation at a lower yield pressure. Another parameter, Da (total degree of densification) for all agglomerates, computed from Equation (4), showed a similar value of 0.603–0.537; this was due to the same particle size range of the agglomerates. The Kawakita plots of the agglomerates are displayed in Figure 3b, and the parameters computed from the relationship between P/C and compression pressure with R2 > 0.99 (compression pressure 31.2–187.3 MPa) are listed in Table 2. The a value, total decrease in powder volume, of the agglomerates showed a similar value in the range of 0.68–0.69. The Pk (yield strength) decreased as the % GTS used for agglomeration increased. This result suggested that agglomerates with a higher % GTS presented lower resistance to volume change under pressure, leading to lower pressure required for particle deformation. In this study, the hardness of the compressed tablets was measured and converted to tensile strength. This parameter related to the compression pressure is shown in Figure 3c. It can be seen that the tablets’ tensile strength increased with increasing pressure. Moreover, the agglomerated tablets without GTS had the lowest tensile strength, and the greater the GTS used, the higher the tablets’ tensile strength was. It is noted that these experiments have a relatively small sample size (n = 3 per compression pressure) and a limited linear range on the Heckel plot (31.2–93.7 MPa).
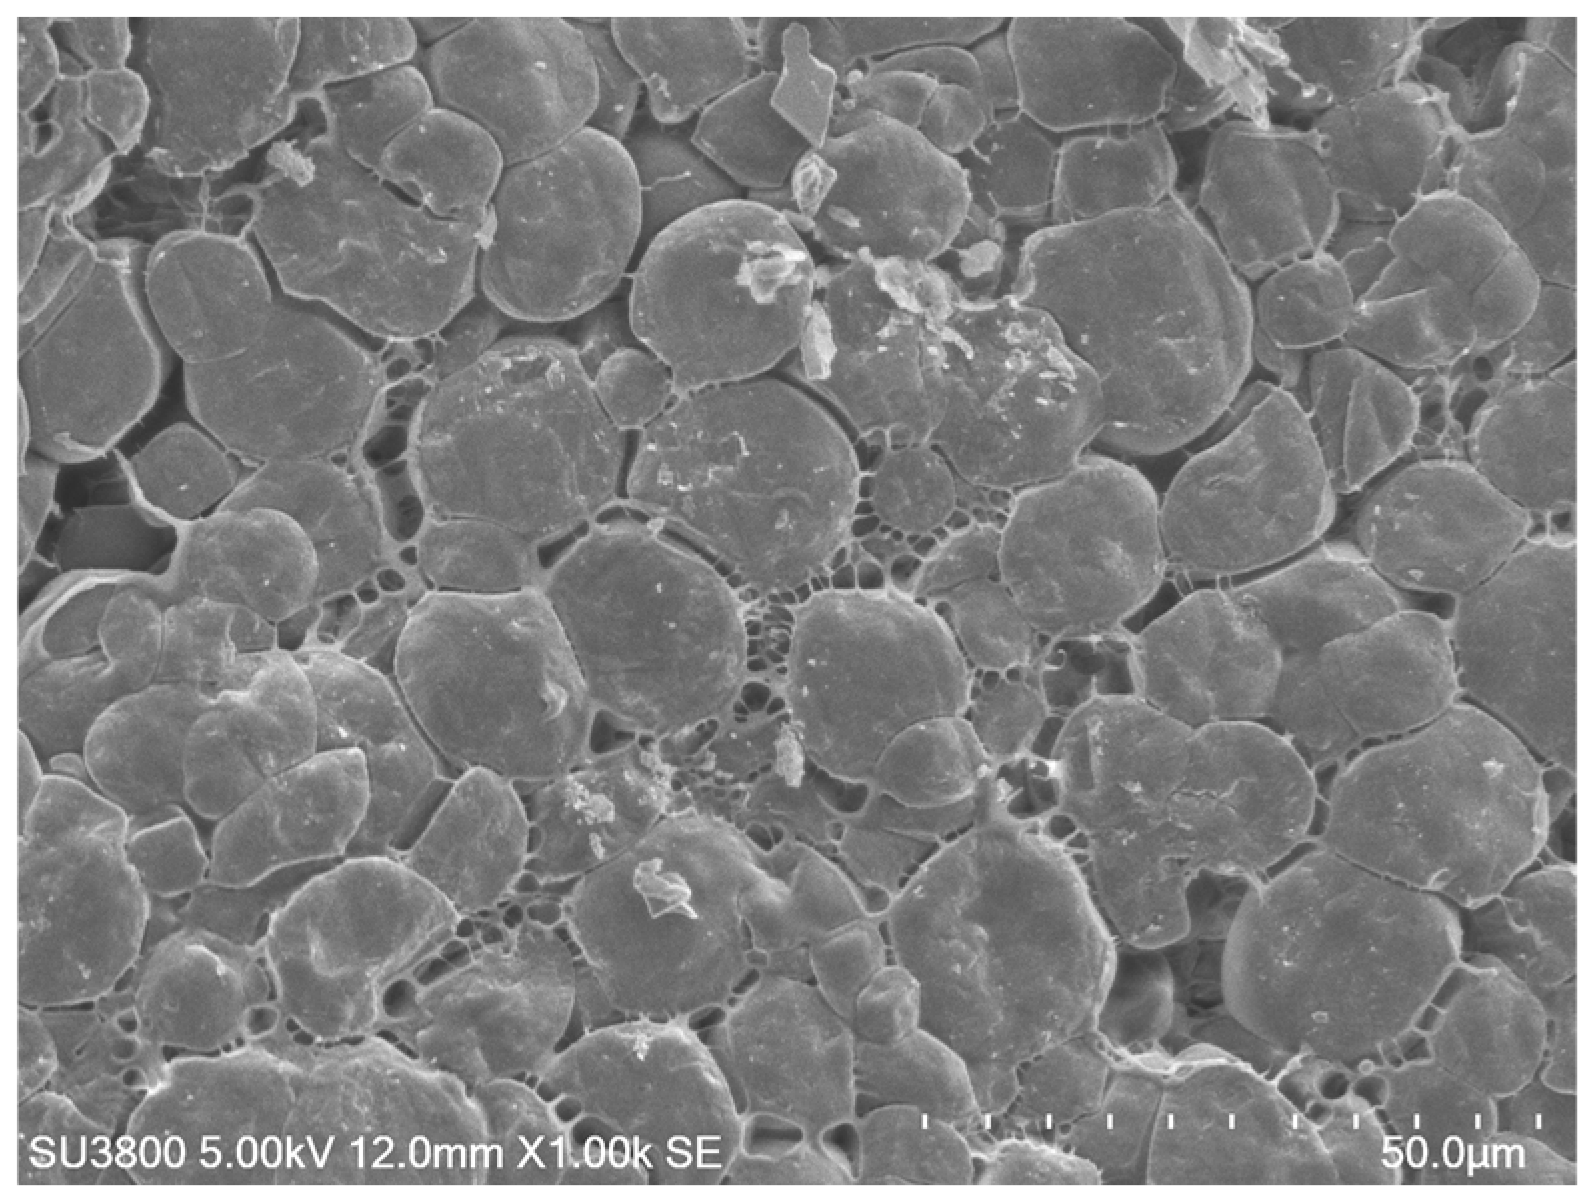
It is well known that starch exhibits plastic deformation under compression pressure [35,36]. To prove this point, the surface morphology of the tablet prepared using TS agglomerates with 2%GTS was observed using SEM, as shown in Figure 4. It can be seen that the starch granules were deformed without fragmentation under compression, and the GTS coated on the surface of TS granules could form a solid bridge with interparticle bonding. Therefore, incorporating GTS as an agglomerating agent could modulate starch particle deformation under pressure. Increasing GTS decreased the yield strength (Pk) and yield pressure (Py) of the agglomerates under pressure, suggesting that agglomerates with higher GTS showed lower resistance to volume reduction and greater particle deformation plasticity. This outcome occurred because the GTS coating on the starch granules could promote plasticity and cold-welding after deformation; thus, agglomerate tablets with higher GTS showed greater tensile strength than those with lower GTS.

3.4. Comparison of Agglomerates Using GTS and Other Materials

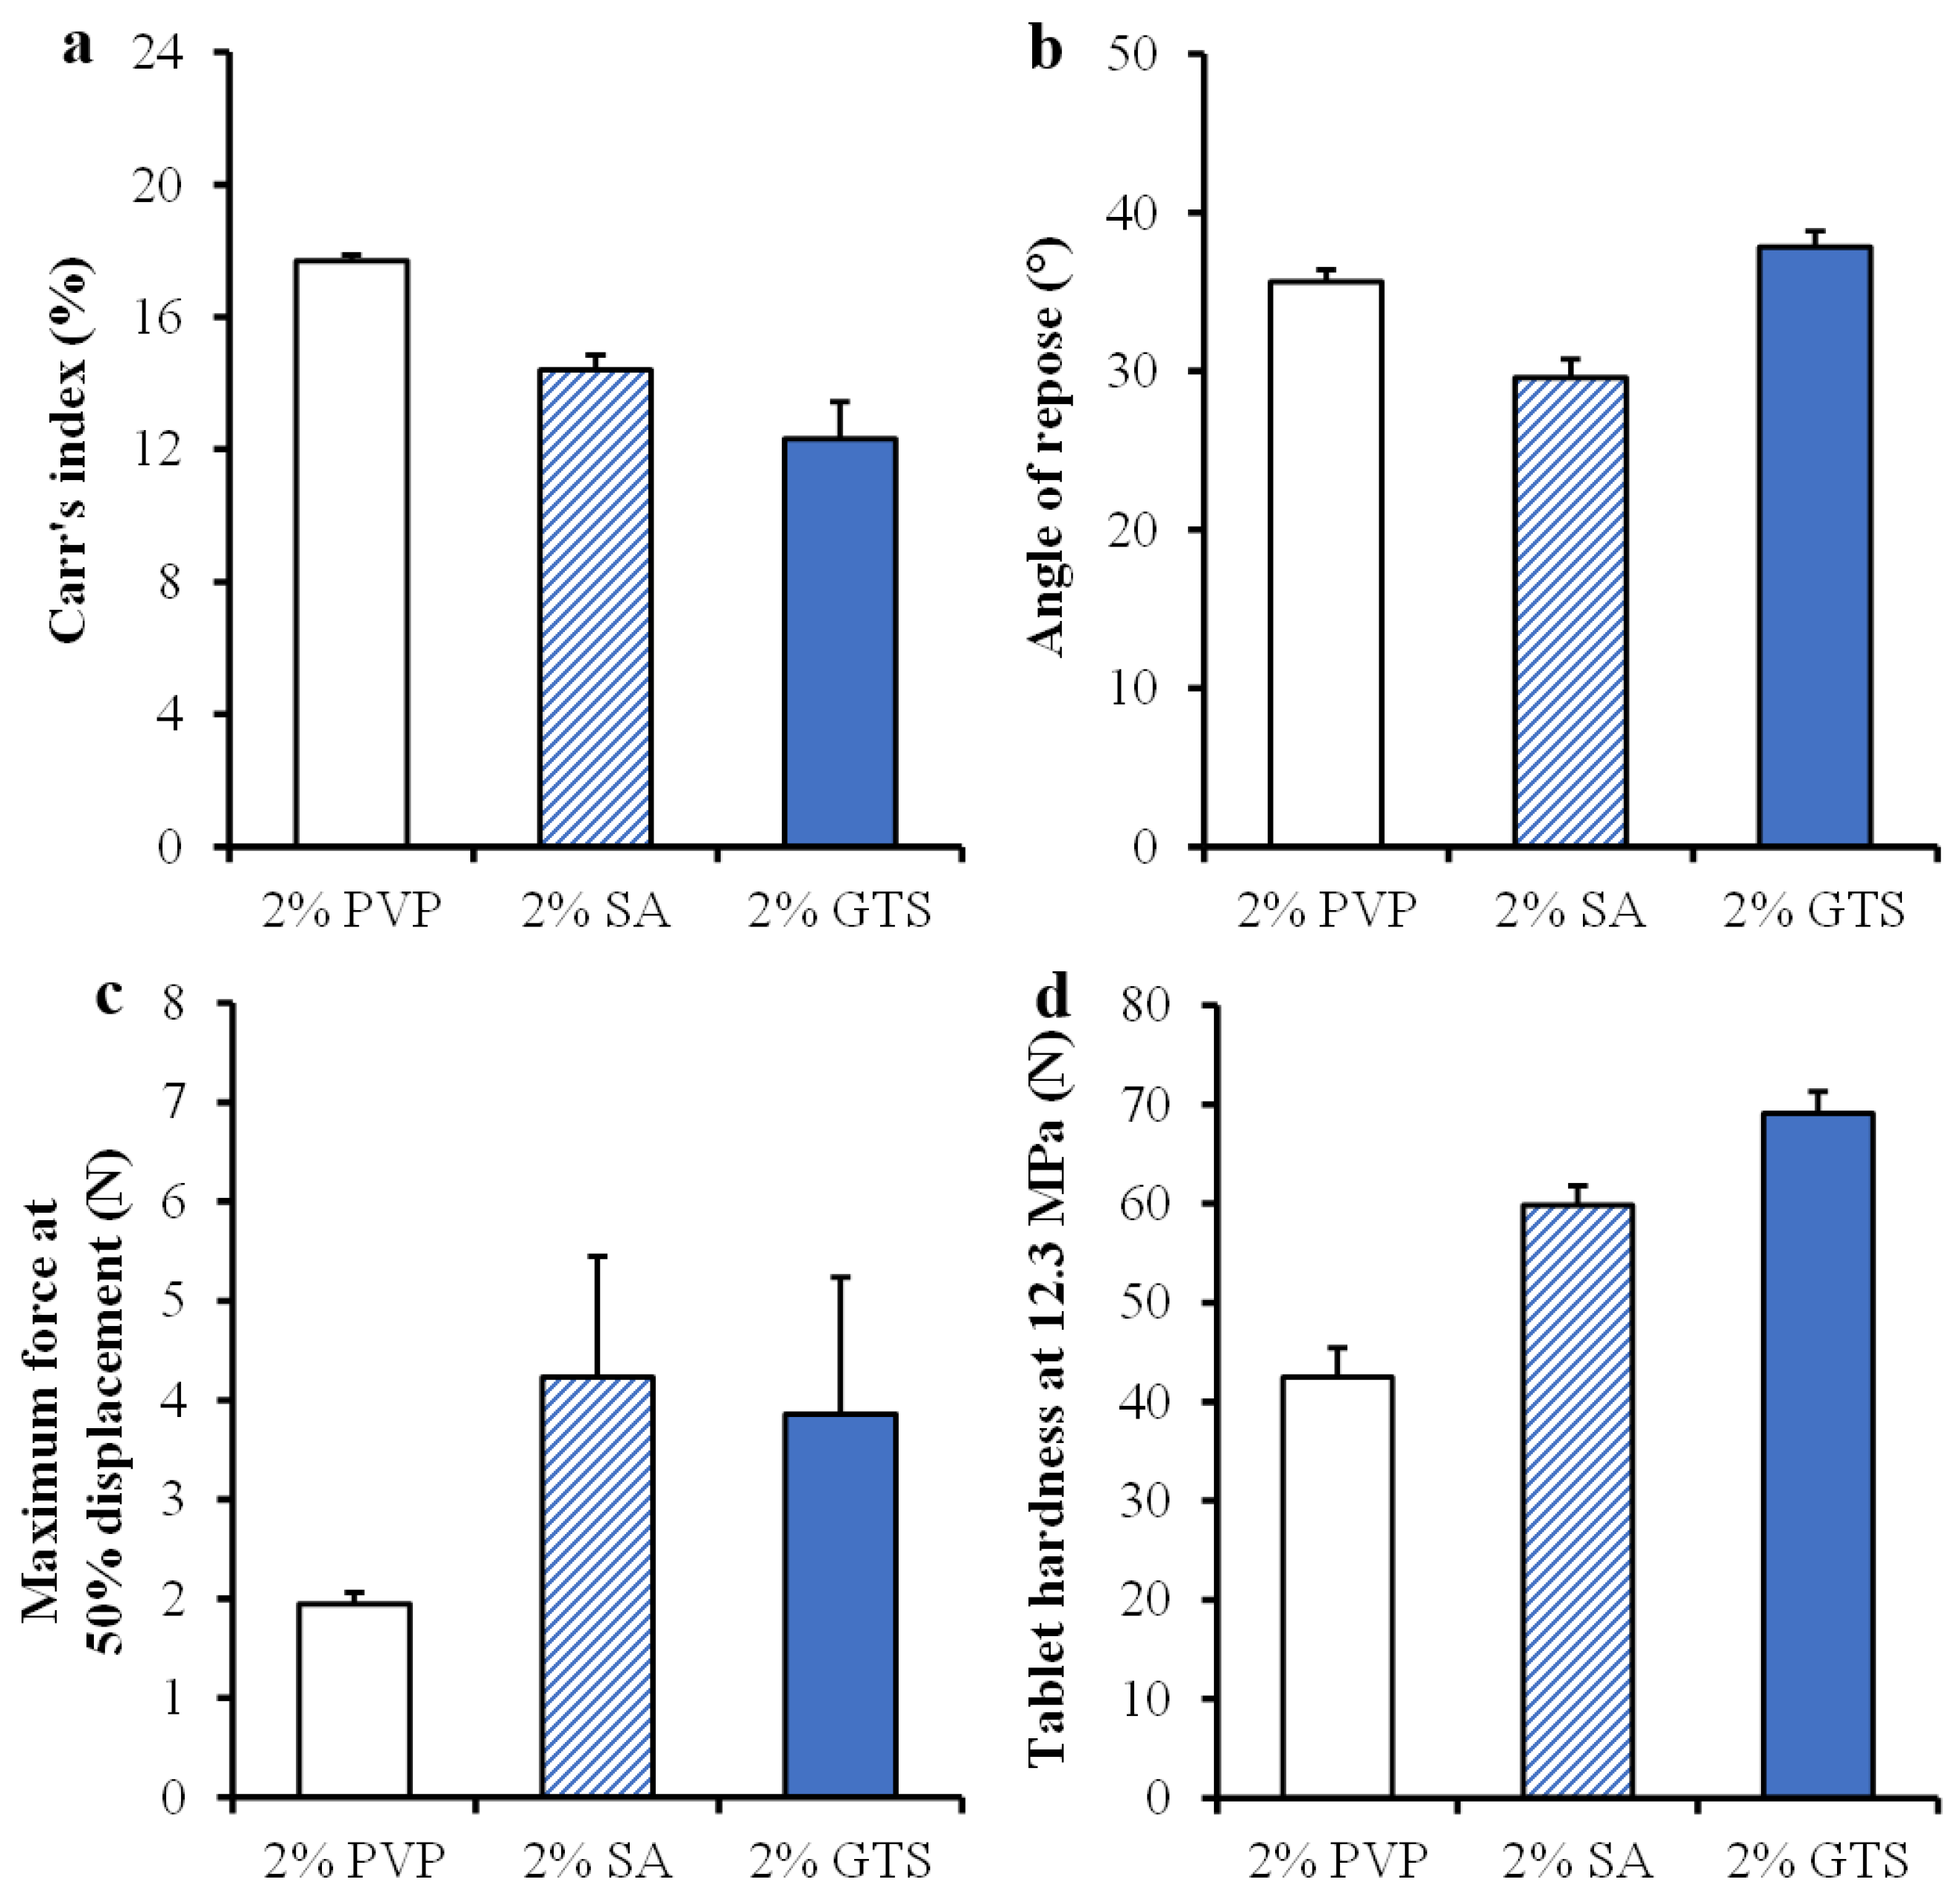
The native TS has been agglomerated using PVP K30 and SA without adding colloidal silicon dioxide, as reported in previous studies [12,20]. In this section, we compare the micromeritic characteristics of TS agglomerates produced using PVP K30 and SA (which has a viscosity of 4–12 cps at 25 °C in a 1% dispersion) with those using GTS at a concentration of 2% w/w. All agglomerates were prepared following the same procedures. The results of Carr’s index and repose angle for the agglomerates are illustrated in Figure 5a,b. These findings indicate that the agglomerates made with the three materials exhibited fair to good flowability, which was an improvement over the native TS. The strength of the agglomerates made with GTS and SA was comparable and superior to that made with PVP K30, as shown in Figure 5c. Moreover, the tablet hardness of the agglomerates produced with GTS (compressed using 12.3 MPa) was greater than that of those made with SA, while the lowest tablet hardness was observed in the agglomerates using PVP K30 (Figure 5d). This comparison demonstrates that particle enlargement through agglomerating agents can enhance the flowability of native TS. However, the strength of the agglomerates and the tablet hardness are influenced by the properties of the materials used. Notably, GTS provided superior compressibility for the agglomerates compared to both PVP K30 and SA. Nevertheless, the presence of colloidal silicon dioxide in the agglomerates using GTS may be limited in this comparison.

3.5. Characteristics of Agglomerate Tablets Loading PNL

The characteristics of agglomerate tablets containing 40 mg of PNL compressed at 12.3 MPa are reported in Table 3. The thickness of the PNL tablets ranged from 2.62 to 2.75 mm (Table 3), whereas those without PNL ranged from 2.71 to 2.77 mm. The hardness of the tablets without PNL was 31.7 ± 1.3, 47.2 ± 1.3, 69.1 ± 2.2, 80.9 ± 2.4, and 96.3 ± 4.6 N (n = 6) when using the agglomerates with 0, 1, 2, 3, and 5% GTS, respectively. These values were statistically higher (p < 0.05) than those of the PNL tablets prepared using the agglomerates with the same GTS content (Table 3). This reduction in hardness is attributed to the interference of drug particles with the cold-welding process of the agglomerates during compression, which deteriorated tablet hardness. However, agglomerates with 0% GTS exhibited the lowest hardness among PNL tablets, whereas increasing GTS content increased hardness. Nevertheless, the PNL tablets containing agglomerates with 3% GTS had hardness comparable to that of those with 5% GTS (Table 3).
The PNL tablets demonstrated rapid disintegration, occurring within 1 min in 0.1 N HCl, as summarized in Table 3. The disintegration time of the PNL tablets without GTS was significantly longer (p < 0.05) than that with GTS, and this parameter decreased with increasing GTS content. However, the PNL tablets with 5% GTS showed no significant difference in disintegration time (p > 0.05) compared with those with 2% or 3% GTS. The results indicate that the rapid disintegration is attributable to the water-absorption properties of TS, which act as a disintegrant in plain tablets [37,38]. As GTS levels in the agglomerates increased, disintegration time decreased in acidic medium (Table 3). As GTS levels in the agglomerates increased, disintegration time decreased in acidic medium (Table 3). This effect is likely due to GTS, a hydrophilic polymer that hydrates and swells in water [37], thereby accelerating tablet disintegration. Moreover, it was observed that increasing GTS content resulted in a shorter disintegration time, whereas tablet hardness increased. Higher GTS content enhanced interparticle bonding through plastic deformation and cold welding within the agglomerates, leading to greater tablet hardness at the same compression pressure. Upon exposure to water, water molecules penetrated the tablet via pore channels. Increased GTS content facilitated greater water absorption, resulting in faster swelling of the gelatinized starch and consequently a shorter disintegration time.
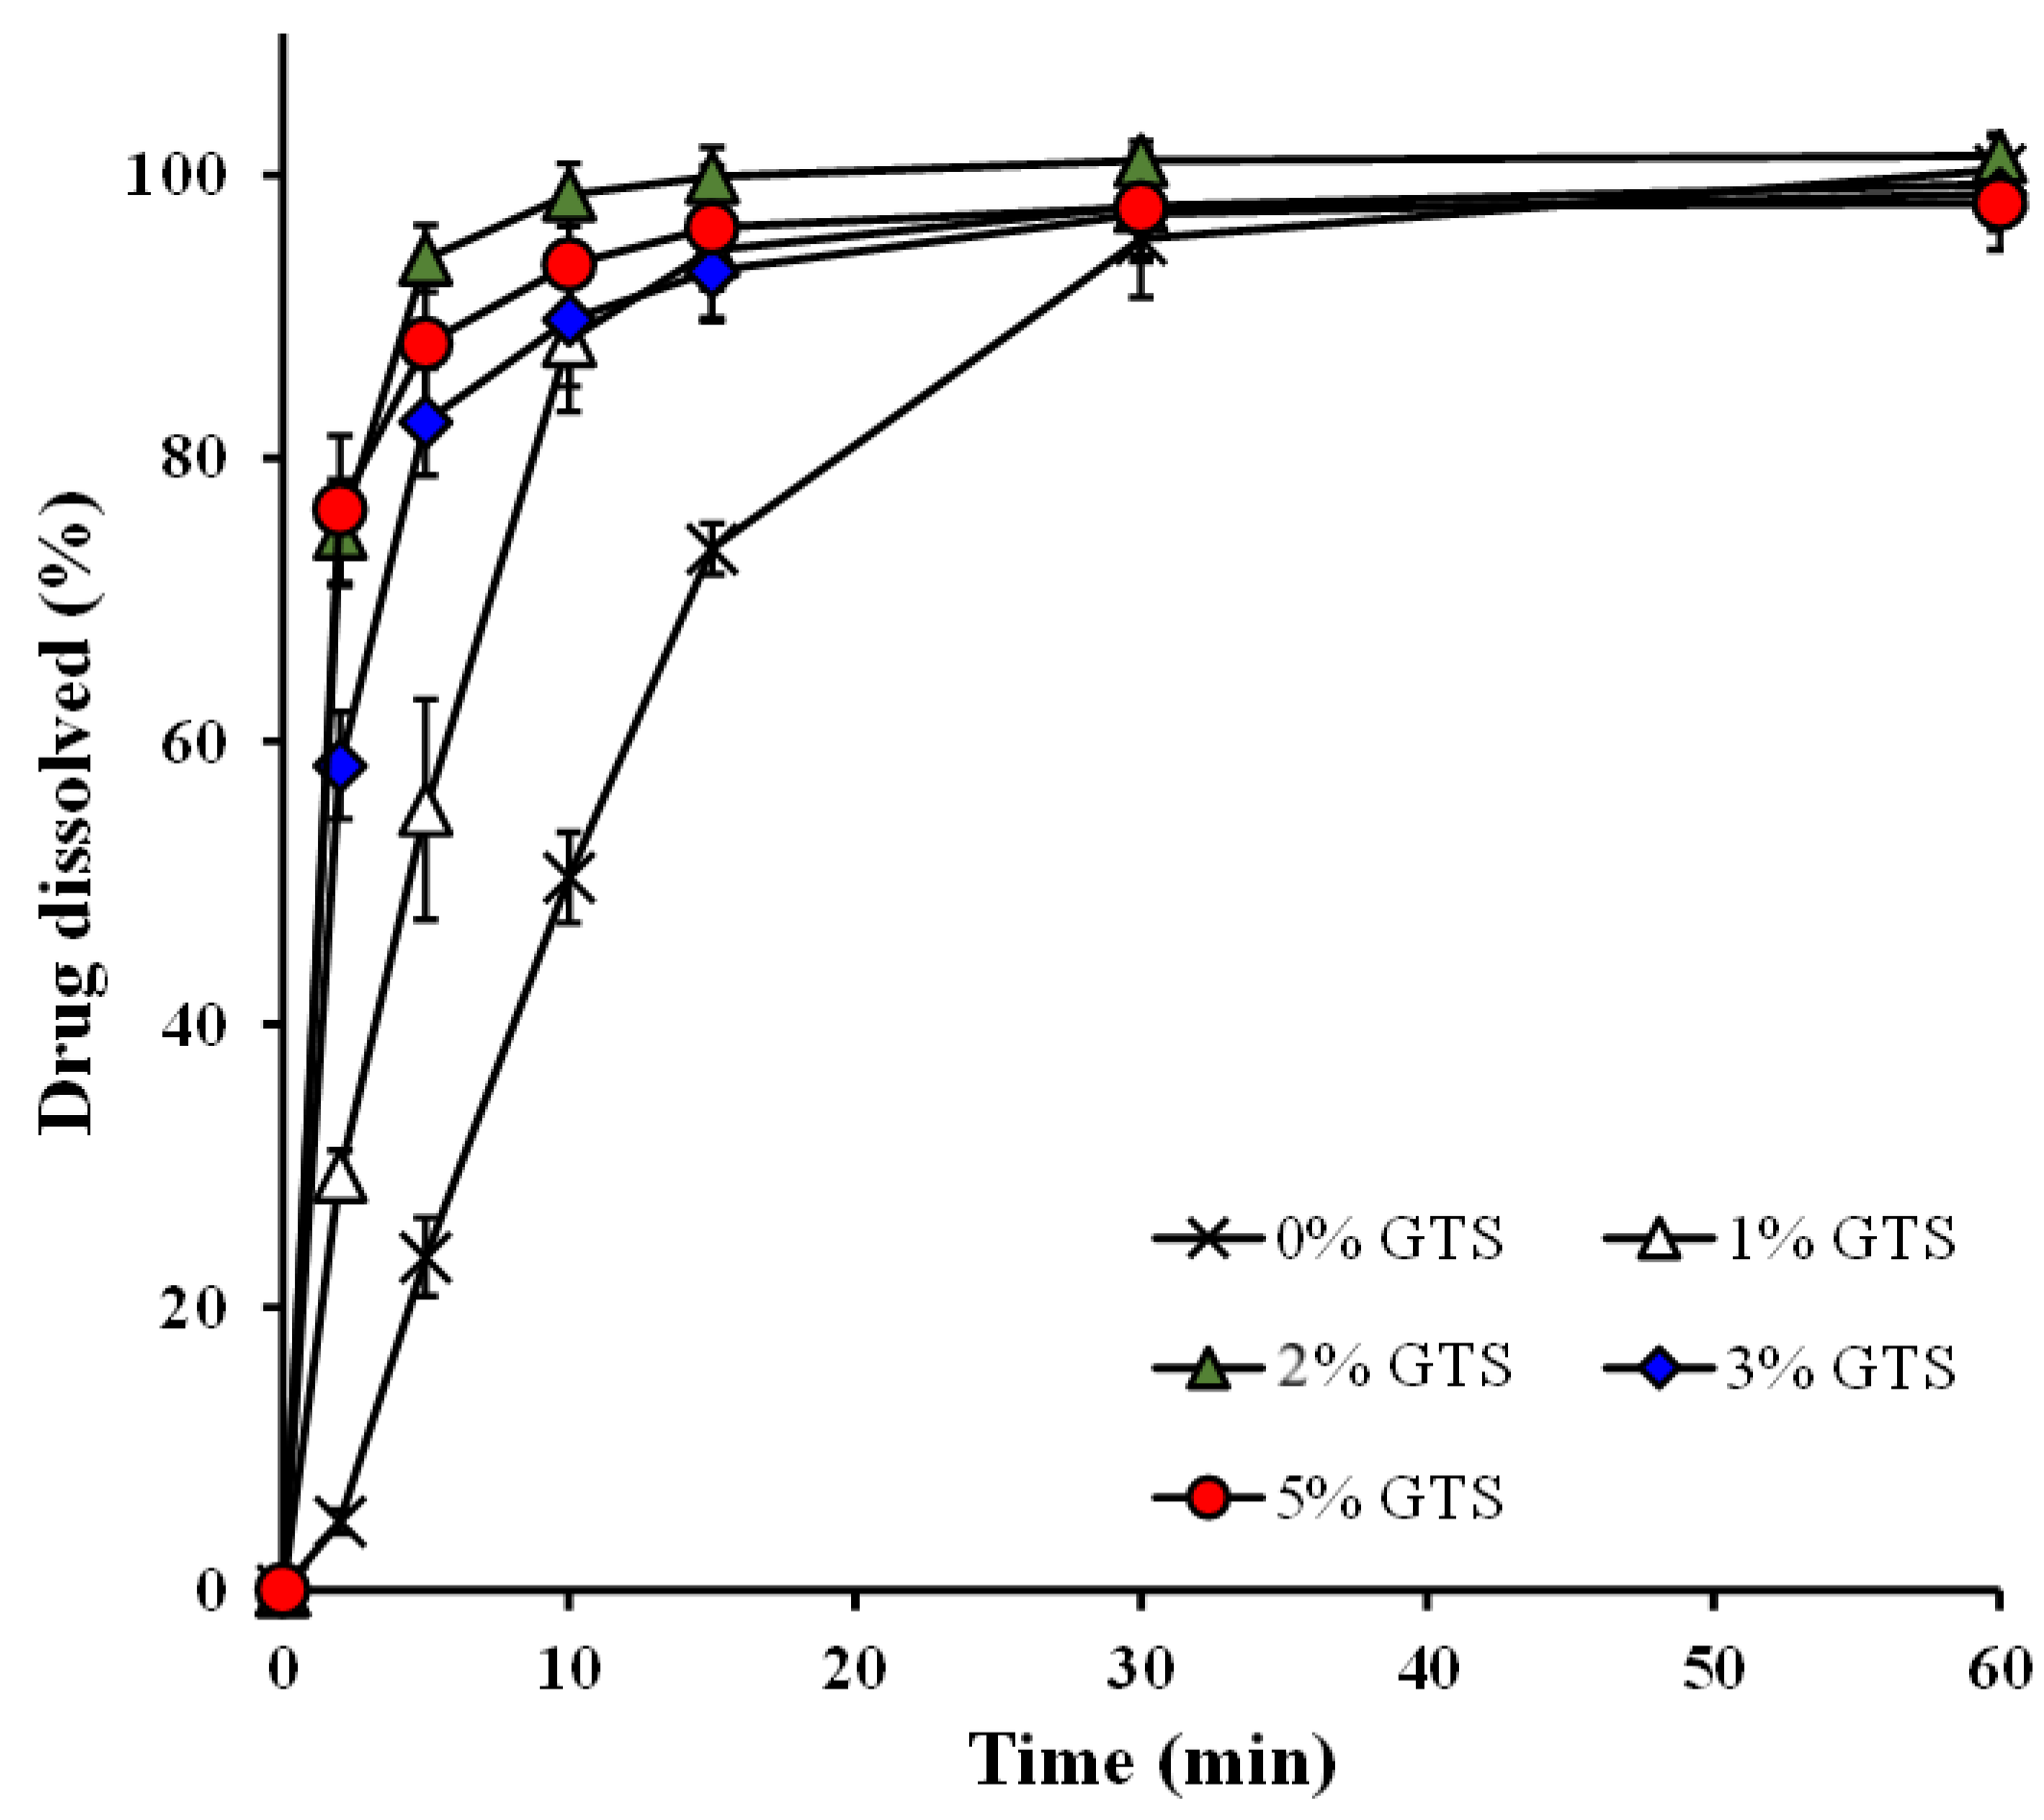
The PNL dissolution profiles of the agglomerate tablets are presented in Figure 6. From the disintegration time obtained, these tablets tended to exhibit an immediate-release dosage form. Thus, the selected medium was 0.1 N HCl to mimic gastric conditions. The result showed that all tablets exhibited rapid drug dissolution, achieving over 85% dissolution within 30 min when tested in 0.1 N HCl. A higher GTS concentration corresponded to a lower T50% value, as shown in Table 3. Additionally, tablets made from agglomerates containing 0% GTS exhibited significantly slower drug dissolution (higher T50%, p < 0.05) than those made with higher GTS content. This finding suggests that GTS enhances drug dissolution from tablets, consistent with the disintegration time results. Overall, GTS appears to improve the properties of the agglomerate tablets by increasing hardness, reducing disintegration time, and accelerating drug dissolution.

3.6. Characteristics of Tablets with Various PNL Loadings

In this study, TS agglomerates prepared with 3% GTS and without colloidal silicon dioxide were used. The moisture content of the agglomerates obtained was 6.89 ± 0.13% w/w (n = 3). The drug loading in tablets was also known as the carrying capacity or dilution potential of the diluents. This capacity was characterized by the highest percentage of non-compressible drugs that could be mixed with the agglomerates to ensure free flow of the mixtures into a die for compression, resulting in acceptable tablet characteristics [39,40]. This property is also referred to as dilution potential. PNL was chosen as the active pharmaceutical ingredient, with a Carr’s index of 34.7 ± 2.2% and a repose angle of 51.9 ± 0.2 degrees (n = 3), indicating poor flowability. Furthermore, pure PNL powder could not form an acceptable tablet upon compression [20], leading to its use as a model drug in this study. The characteristics of agglomerate tablets with different PNL loadings are detailed in Table 4. The tablets were approximately 3.5 mm thick and weighed 348.3–355.9 mg. The relative standard deviation (RSD) of the tablet weights tended to increase with higher PNL loadings. This result suggests that a greater percentage of PNL loading affected the flow of the mixtures into the die prior to compression. However, the % RSD of tablet weight was 0.54 and 0.95 at 30 and 40% PNL loading, respectively, suggesting good flowability of the mixtures when using agglomerates as a diluent. This is why the PNL content in the tablets remained consistent at 96.6–101.9%.
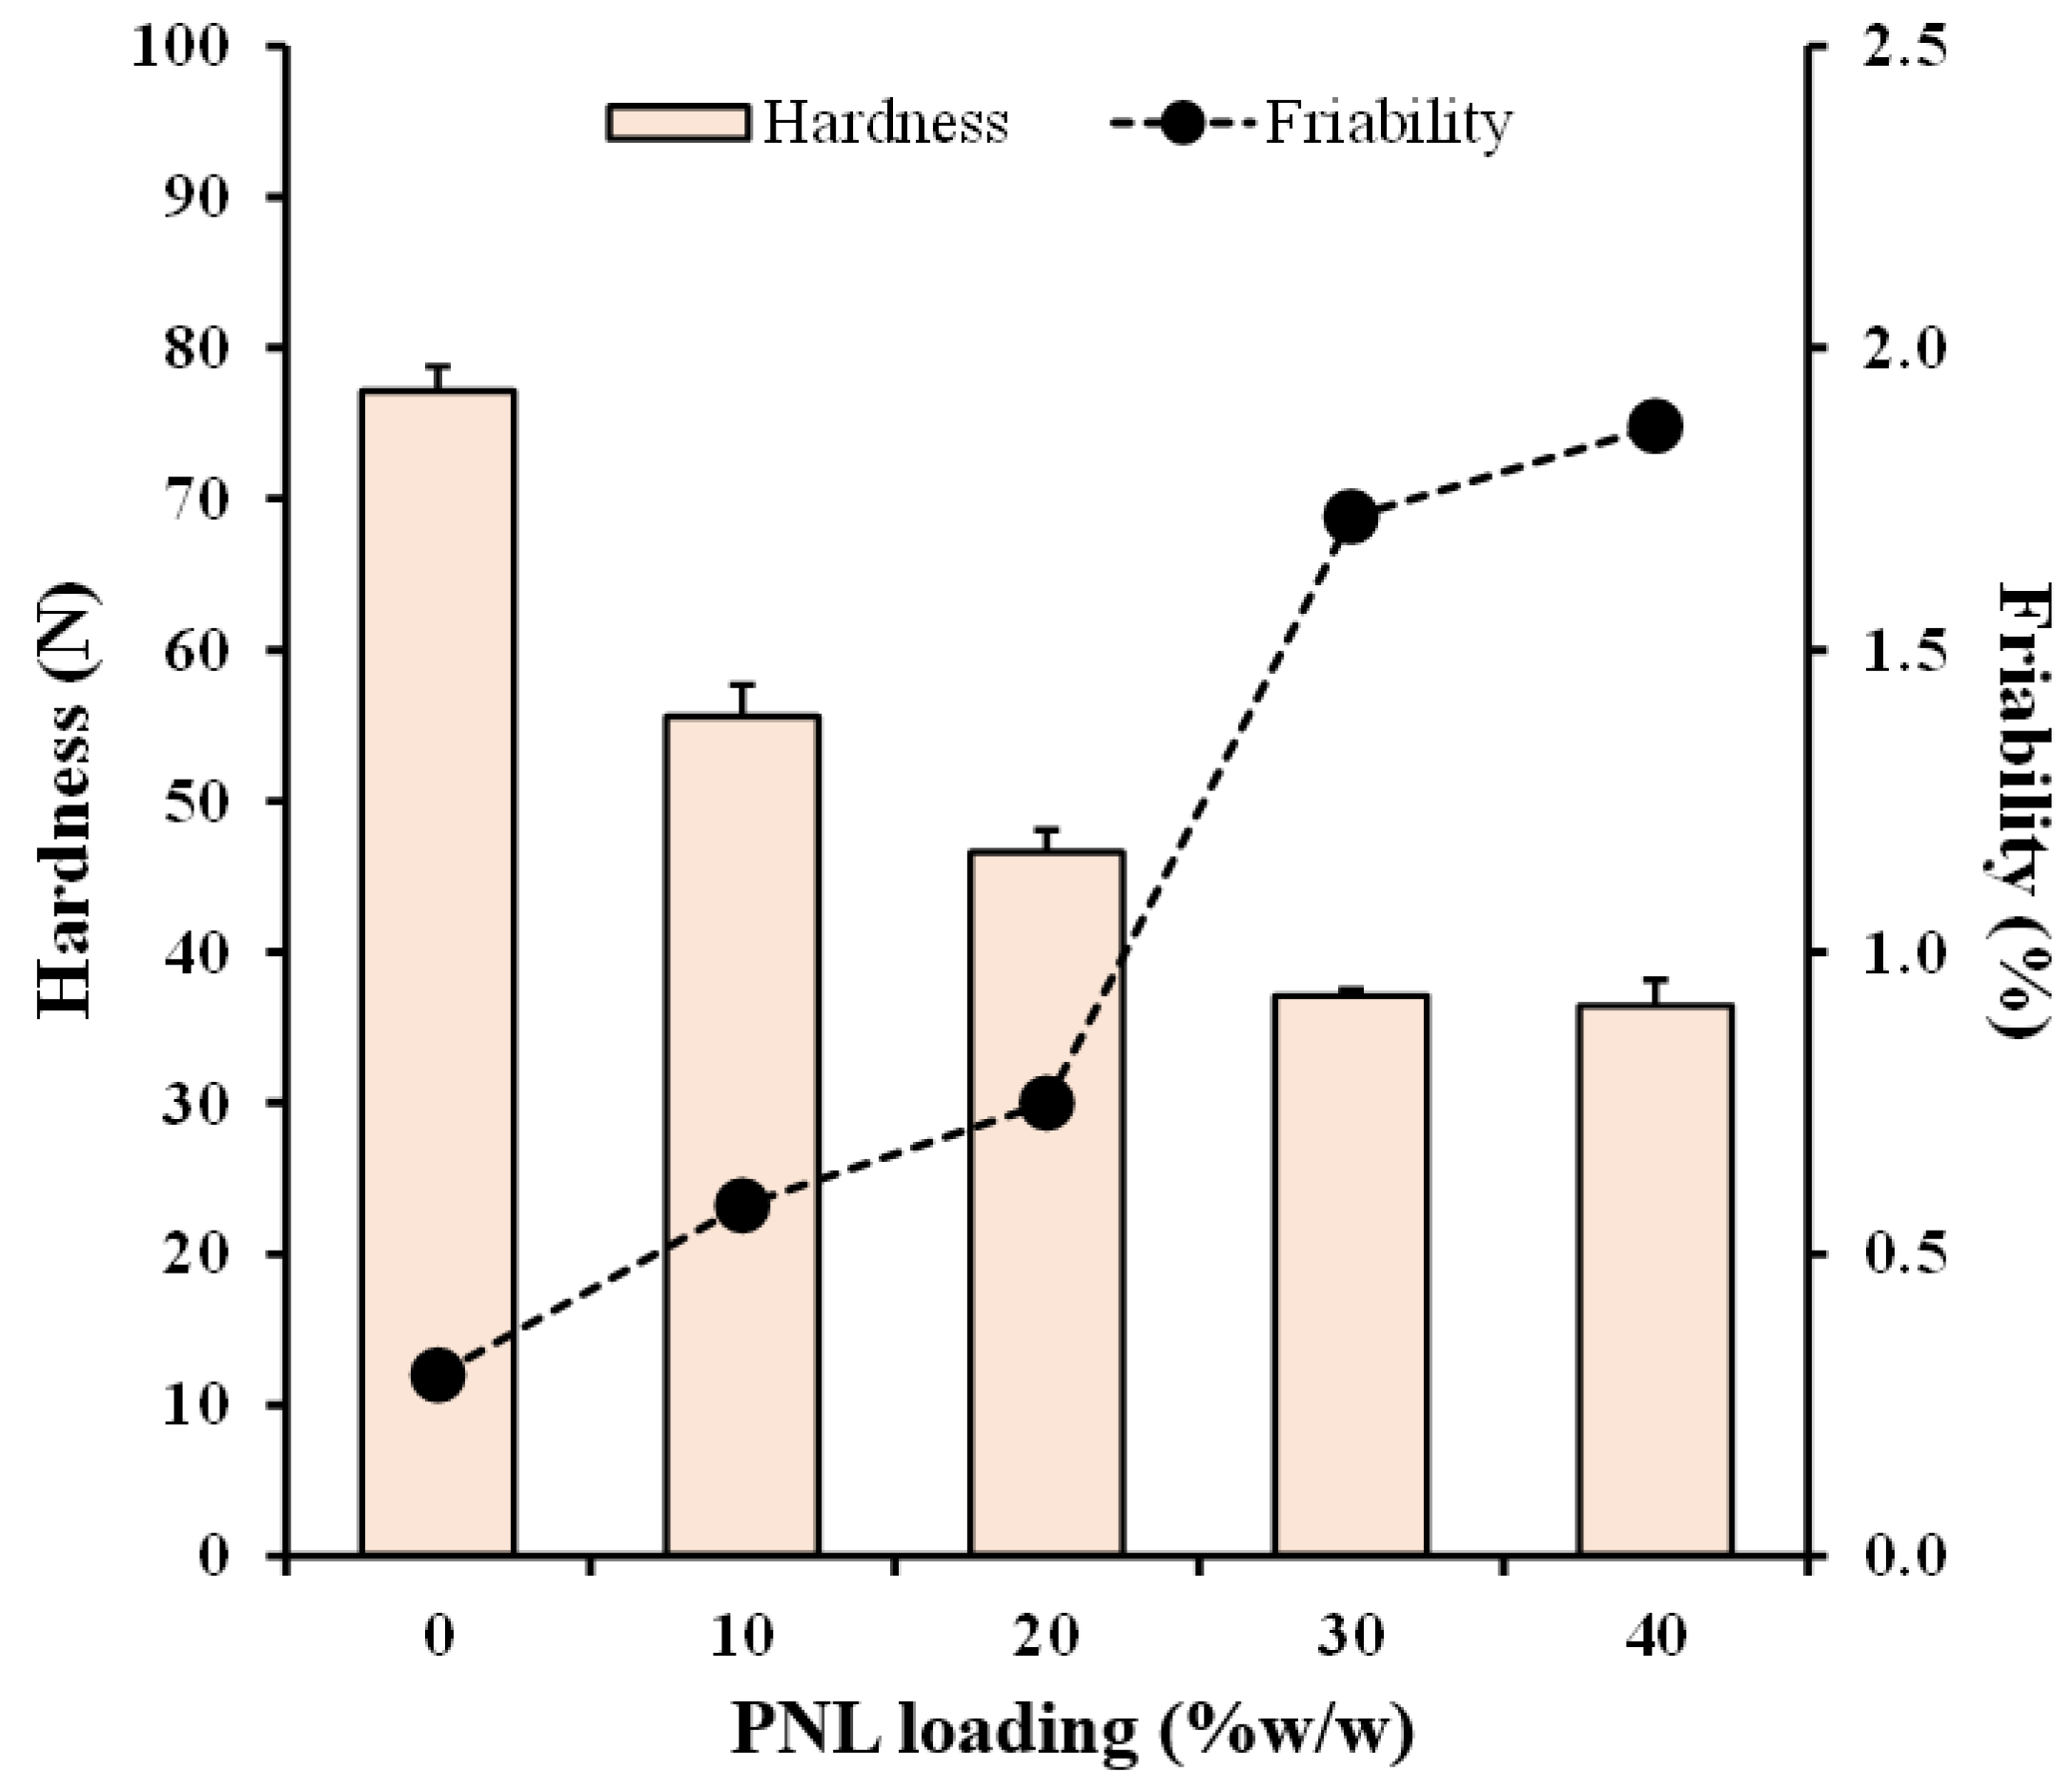
Both the hardness and friability of the tablets were influenced by the PNL loading percentage, as shown in Figure 7. An increase in PNL loading resulted in reduced tablet hardness and increased friability. This outcome was attributed to the disturbance of the cold-welding process of the agglomerates caused by the incorporation of PNL. The tablets loaded with 0–20% w/w PNL showed a friability value less than 1% (n = 1; this parameter was tested from an average weight loss of the tablets not less than 6.5 g), and an increase in PNL loadings to 30 and 40% w/w resulted in a friability value greater than unity (1.72 and 1.87%, respectively). Moreover, these PNL loadings resulted in tablet hardness dropping to 37.10 ± 0.40 and 36.45 ± 1.69 N (n = 6), respectively. A common criterion for tablet friability was a weight loss of less than 1.0% after testing using the method in Section 2.5, which was a requirement in the US Pharmacopeia [41]. Therefore, tablets with 30% and 40% PNL loading did not meet this criterion due to chipping of the tablet surface and edges during mechanical rotation. In contrast, tablet hardness, defined as the force required to break diametrically, does not have a specified threshold in the US Pharmacopeia. Despite reduced hardness, tablets at these PNL loadings retained sufficient structural integrity. Consequently, friability, coupled with tablet hardness, should be considered in the early stages of tablet development [41]. To address the observed deficiencies, increasing compression pressure for tablets with higher PNL loading (30–40% PNL), which demonstrate good flowability and PNL content, may enhance both hardness and friability.
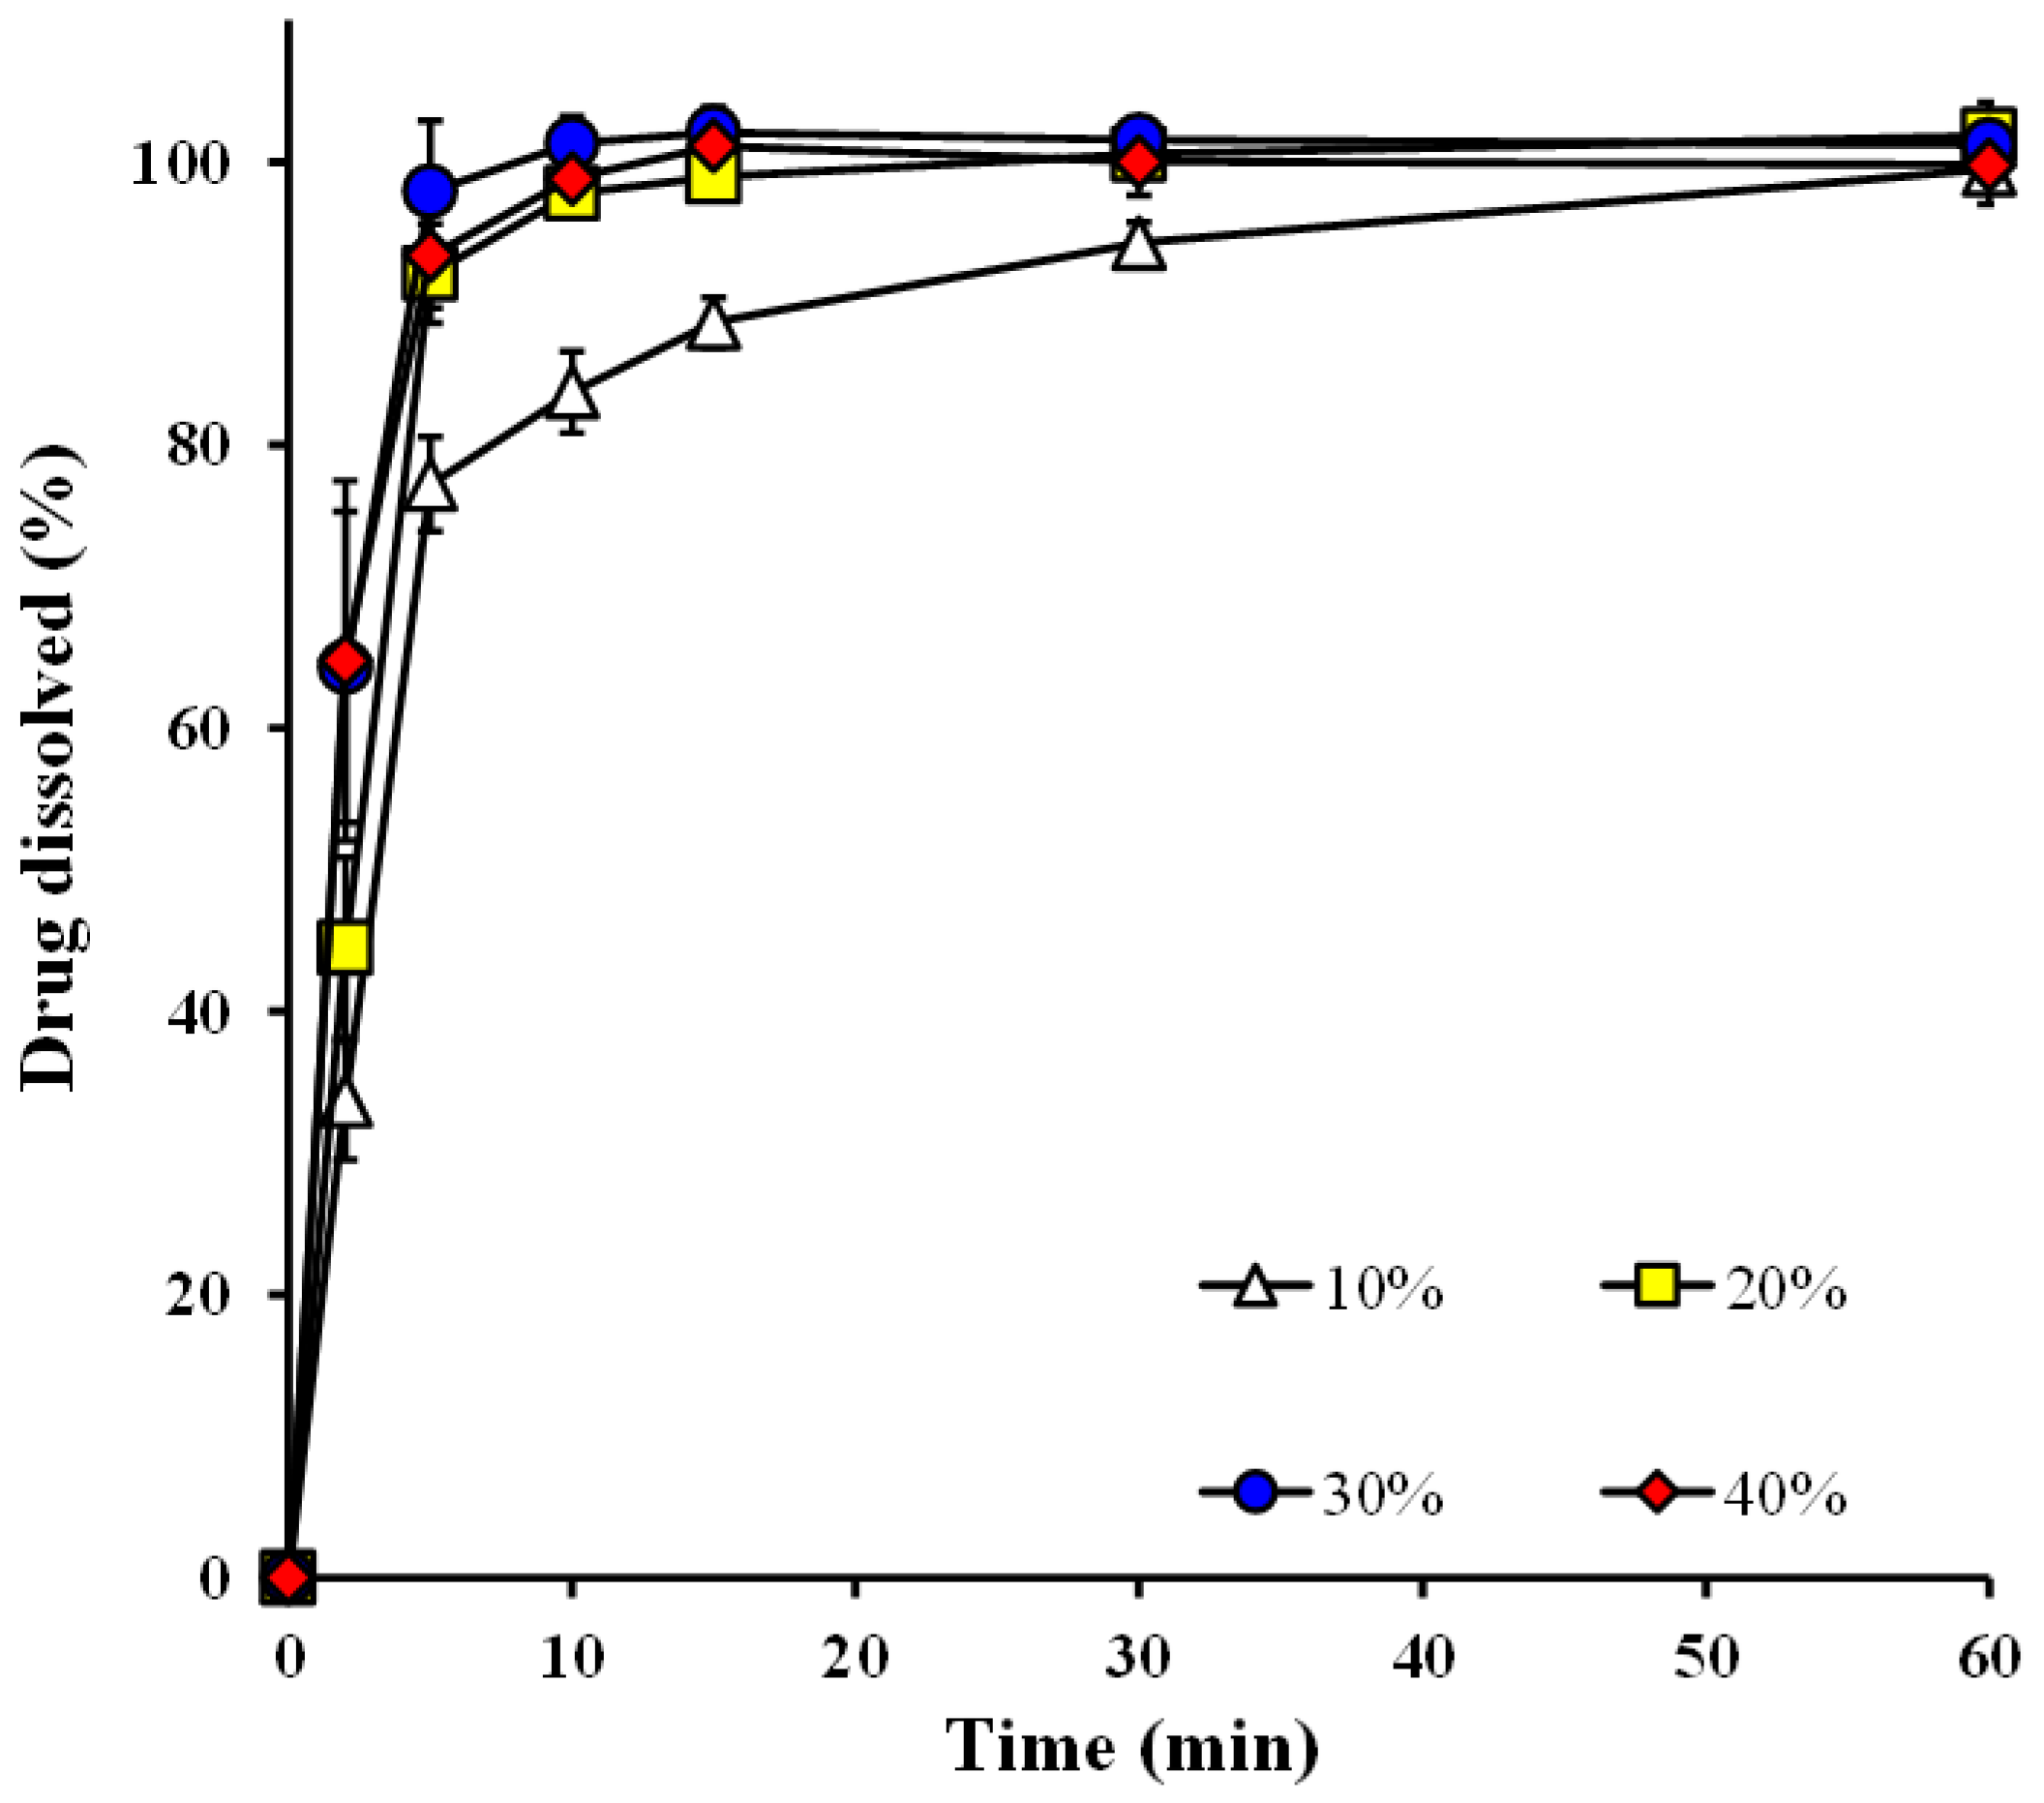
The disintegration time in an acidic medium decreased with increasing drug loading (Table 4), attributed to decreased tablet hardness. The tablets also displayed rapid drug dissolution in the acidic medium, as shown in Figure 8. The T50% value tended to decrease with increasing PNL loading percentage (Table 4). According to the requirements of the Brazilian and US Pharmacopeias, PNL tablets were recommended to achieve at least 75% drug dissolution within 30 min using dissolution apparatus 1, with a medium of 1000 mL of 0.1 N HCl and a basket speed of 100 revolutions/min [42]. The tablets in this study gave more than 90% PNL dissolution within 30 min. Notably, the volume of the dissolution medium (750 mL) and the basket rotation speed (50 revolutions/min) used in this study were lower than the conditions in the pharmacopeia. This result suggested that the PNL tablets exhibited rapid disintegration and an immediate-release profile, which met the requirements of the pharmacopeia.
This study used a single water-soluble PNL with low flowability and compressibility. Additional active ingredients, including poorly water-soluble drugs, high-dose drugs, highly elastic drugs, and crude herbal powders, should be evaluated for use with TS agglomerates containing GTS. Preliminary testing of these formulations is recommended.

4. Conclusions

This study revealed the role of GTS as an agglomerating agent in producing TS agglomerates. Enlargement of TS granules using GTS resulted in an increase in agglomerates’ strength and better flowability of the agglomerates compared to that of native TS, as indicated by Carr’s index and repose angle. In addition, the compressibility of the TS agglomerates increased with increasing GTS content, leading to lower resistance to volume reduction and greater plasticity of particle deformation under pressure, resulting in a greater tensile strength of the tablets. The PNL tablets made from agglomerates with GTS had higher hardness than those without GTS, and increasing GTS content increased tablet hardness. Moreover, GTS could accelerate tablet disintegration, resulting in faster PNL dissolution from the tablets. The agglomerates with 3% GTS were used to produce PNL tablets for a carrying capacity or dilution potential test. The agglomerate tablets exhibited acceptable physical properties when the PNL loading was less than 20% w/w in this study. These results suggest that native TS agglomerates produced with GTS could serve as a tablet diluent for direct compression. However, the tablets with drug loadings of 30% and 40% require further investigation under varying compression pressures.

Author Contributions

Conceptualization, T.P.; methodology, R.J. and C.S.; validation, T.P.; formal analysis, R.J. and C.S.; investigation, R.J. and C.S.; data curation, R.J.; writing—original draft preparation, R.J.; writing—review and editing, T.P.; visualization, T.P.; supervision, T.P. All authors have read and agreed to the published version of the manuscript.

Funding

This research received no external funding.

Institutional Review Board Statement

Not applicable.

Informed Consent Statement

Not applicable.

Data Availability Statement

The original contributions presented in this study are included in the article. Further inquiries can be directed to the corresponding author.

Acknowledgments

The authors would like to thank the Faculty of Pharmaceutical Sciences and Center for Research and Development of Herbal Health Products, Khon Kaen University (Khon Kaen, Thailand) for technical supports.

Conflicts of Interest

The authors declare no conflicts of interest.

References

  1. Charoenthai, N.; Sanga-ngam, T.; Puttipipatkhachorn, S. Use of modified tapioca starches as pharmaceutical excipients. Pharm. Sci. Asia 2018, 45, 195–204. [Google Scholar] [CrossRef] [Scilit]
  2. Elgaied-Lamouchi, D.; Descamps, N.; Lefevre, P.; Rambur, I.; Pierquin, J.Y.; Siepmann, F.; Siepmann, J.; Muschert, S. Starch-based controlled release matrix tablets: Impact of the type of starch. J. Drug Deliv. Sci. Technol. 2021, 61, 102152. [Google Scholar] [CrossRef] [Scilit]
  3. Das, S.; Das, M.K.; Jamatia, T.; Bhattacharya, B.; Mazumder, R.; Yadav, P.K.; Bishwas, N.R.G.; Deka, T.; Roy, D.; Sinha, B.; et al. Advances of cassava starch-based composites in novel and conventional drug delivery systems: A state-of-the-art review. RSC Pharm. 2024, 1, 182–203. [Google Scholar] [CrossRef] [Scilit]
  4. Zarski, A.; Kapusniak, K.; Ptak, S.; Rudlicka, M.; Coseri, S.; Kapusniak, J. Functionalization methods of starch and its derivatives: From old limitations to new possibilities. Polymers 2024, 16, 597. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  5. Yamini, K.; Chalapathi, V.; Lakshmi Narasimha Reddy, N.; Lokesh, K.V.; Praveen Kumar Reddy, S.; Gopal, V. Formulation of diclofenac sodium tablets using tapioca starch powder-A promising binder. J. Appl. Pharm. Sci. 2011, 1, 125–127. [Google Scholar]
  6. Mishra, S.; Rai, T. Morphology and functional properties of corn, potato and tapioca starches. Food Hydrocoll. 2006, 20, 557–566. [Google Scholar] [CrossRef] [Scilit]
  7. Sanguanpong, V.; Chotineeranat, S.; Piyachomkwan, K.; Oates, C.G.; Chinachoti, P.; Sriroth, K. Hydration and physicochemical properties of small-particle cassava starch. J. Sci. Food Agric. 2003, 83, 123–132. [Google Scholar] [CrossRef] [Scilit]
  8. Siriwachirachai, C.; Pongjanyakul, T. Acid and alkali modifications of tapioca starches: Physicochemical characterizations and evaluations for use in tablets. J. Drug Deliv. Sci. Technol. 2022, 68, 103068. [Google Scholar] [CrossRef] [Scilit]
  9. Chitedze, J.; Monjerezi, M.; Saka, J.D.K.; Steenkamp, J. Binding effect of cassava starches on the compression and mechanical properties of ibuprofen tablets. J. Appl. Pharm. Sci. 2012, 2, 31–37. [Google Scholar] [CrossRef] [Scilit]
  10. Lawal, M.V. Modified starches as direct compression excipients–Effect of physical and chemical modifications on tablet properties: A review. Starch-Stärke 2019, 71, 1800040. [Google Scholar] [CrossRef] [Scilit]
  11. Atichokudomchai, N.; Shobsngob, S.; Chinachoti, P.; Varavinit, S. A study of some physicochemical properties of high-crystalline tapioca starch. Starch-Stärke 2001, 53, 577–581. [Google Scholar] [CrossRef] [Scilit]
  12. Siriwachirachai, C.; Pongjanyakul, T. Particle agglomeration of acid-modified tapioca starches: Characterization and use as direct compression fillers in tablets. Pharmaceutics 2022, 14, 1245. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  13. Atichokudomchai, N.; Varavinit, S. Characterization and utilization of acid-modified cross-linked tapioca starch in pharmaceutical tablets. Carbohydr. Polym. 2003, 53, 263–270. [Google Scholar] [CrossRef] [Scilit]
  14. Triwises, P.; Ruksakulpiwat, Y.; Ruksakulpiwat, C. The modification of tapioca starch by esterification technique. Suranaree J. Sci. Technol. 2016, 23, 157–165. [Google Scholar]
  15. Pornpitchanarong, C.; Bangsitthideth, K.; Pengnam, S.; Charoenying, T.; Pamornpathomkul, B.; Taesotikul, T.; Patrojanasophon, P.; Chaksmithanont, P. Optimization of modified taro (Colocasia esculenta) starch as a functional binder for pharmaceutical tablets. J. Drug Deliv. Sci. Technol. 2026, 115, 107710. [Google Scholar] [CrossRef] [Scilit]
  16. Pietsch, W. An interdisciplinary approach to size enlargement by agglomeration. Powder Technol. 2003, 130, 8–13. [Google Scholar] [CrossRef] [Scilit]
  17. Khlibsuwan, R.; Pongjanyakul, T. Particle agglomeration of chitosan–magnesium aluminum silicate nanocomposites for direct compression tablets. Int. J. Pharm. 2018, 535, 410–419. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  18. Jarungsirawat, R.; Kajthunyakarn, W.; Siriwachirachai, C.; Pongjanyakul, T. Formulation development of fluconazole-loaded lactose agglomerate tablets as a disinfectant for Candida-associated dentures. Pharmaceutics 2022, 14, 1723. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  19. Huang, S.; Liu, Y.; Zhou, Y.; Li, Q.; Ren, G.; Jing, Q.; Ren, F. Exploring the effect of PVP on the spherical agglomeration process and micromeritic properties of ascorbic acid. Powder Technol. 2019, 342, 929–937. [Google Scholar] [CrossRef] [Scilit]
  20. Jarungsirawat, R.; Siriwachirachai, C.; Jaipakdee, N.; Chitropas, P.; Pongjanyakul, T. Agglomeration of native tapioca starch using sodium alginate for use in tablets. J. Drug Deliv. Sci. Technol. 2024, 101, 106237. [Google Scholar] [CrossRef] [Scilit]
  21. Siriwachirachai, C.; Pongjanyakul, T. Gelatinized tapioca starch−magnesium aluminum silicate nanocomposites: Physicochemical characterization and evaluation for modified-release tablets. J. Drug Deliv. Sci. Technol. 2023, 85, 104591. [Google Scholar] [CrossRef] [Scilit]
  22. Siriwachirachai, C.; Pongjanyakul, T. Gelatinized tapioca starch-magnesium aluminum silicate nanocomposites: Particle agglomeration for directly compressed tablets. J. Drug Deliv. Sci. Technol. 2023, 87, 104853. [Google Scholar] [CrossRef] [Scilit]
  23. Jarungsirawat, R.; Tabboon, P.; Limpongsa, E.; Sakloetsakun, D.; Pongjanyakul, T.; Jaipakdee, N. Material properties, compressibility, and tabletability of planetary ball-milled glutinous rice starch and its application as a mucoadhesive disc for buccal delivery: Impact of milling duration. Int. J. Biol. Macromol. 2025, 306, 141612. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  24. Sen, O.; Pal, D.; Jana, T.; Das, S.; Nandid, G.; Manna, S. Extraction, characterization and evaluation of jute (Corchorus olitorius) leaf polysaccharide as a binding agent for matrix tablet formulation. RSC Pharm. 2026, 3, 490–506. [Google Scholar] [CrossRef] [Scilit]
  25. Shang, C.; Sinka, I.C.; Jayaraman, B.; Pan, J. Break force and tensile strength relationships for curved faced tablets subject to diametrical compression. Int. J. Pharm. 2013, 442, 57–64. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  26. Yohannes, B.; Abebe, A. Determination of tensile strength of shaped tablets. Powder Technol. 2021, 383, 11–18. [Google Scholar] [CrossRef] [Scilit]
  27. Hassanpour, A.; Ghadiri, M. Distinct element analysis and experimental evaluation of the heckel analysis of bulk powder compression. Powder Technol. 2004, 141, 251–261. [Google Scholar] [CrossRef] [Scilit]
  28. Choi, D.H.; Kim, N.A.; Chu, K.R.; Jung, Y.J.; Yoon, J.H.; Jeong, S.H. Material properties and compressibility using Heckel and Kawakita equation with commonly used pharmaceutical excipients. J. Pharm. Investig. 2010, 40, 237–244. [Google Scholar] [CrossRef] [Scilit]
  29. Khomane, K.S.; Bansal, A.K. Effect of particle size on in-die and out-of-die compaction behavior of ranitidine hydrochloride polymorphs. AAPS Pharm. Sci. Technol. 2013, 14, 1169–1177. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  30. Simons, S.J.R.; Rossetti, D.; Pagliai, P.; Ward, R.; Fitzpatrick, S. The relationship between surface properties and binder performance in granulation. Chem. Eng. Sci. 2005, 60, 4055–4060. [Google Scholar] [CrossRef] [Scilit]
  31. Carr, R.L. Evaluating flow properties of solids. Chem. Eng. 1965, 72, 163–168. [Google Scholar]
  32. Horio, T.; Yasuda, M.; Matsusaka, S. Effect of particle shape on powder flowability of microcrystalline cellulose as determined using the vibration shear tube method. Int. J. Pharm. 2014, 473, 572–578. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  33. Leturia, M.; Benali, M.; Lagarde, S.; Ronga, I.; Saleh, K. Characterization of flow properties of cohesive powders: A comparative study of traditional and new testing methods. Powder Technol. 2014, 253, 406–423. [Google Scholar] [CrossRef] [Scilit]
  34. Kudo, Y.; Yasuda, M.; Matsusaka, S. Effect of particle size distribution on flowability of granulated lactose. Adv. Powder Technol. 2020, 31, 121–127. [Google Scholar] [CrossRef] [Scilit]
  35. Paronen, P.; Juslin, M. Compressional characteristics of four starches. J. Pharm. Pharmacol. 1983, 35, 627–635. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  36. Akin-Ajani, O.D.; Itiola, O.A.; Odeku, O.A. Effect of acid modification on the material and compaction properties of fonio and sweet potato starches. Starch-Stärke 2014, 66, 749–759. [Google Scholar] [CrossRef] [Scilit]
  37. Markl, D.; Zeitler, J.A. A review of disintegration mechanisms and measurement techniques. Pharm. Res. 2017, 34, 890–917. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  38. Desai, P.M.; Liew, C.V.; Heng, P.W.S. Review of disintegrants and the disintegration phenomena. J. Pharm. Sci. 2016, 105, 2545–2555. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  39. Habib, Y.; Augsburger, L.; Reier, G.; Wheatley, T.; Shangraw, R. Dilution potential: A new perspective. Pharm. Dev. Technol. 1996, 1, 205–212. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  40. Mitrevej, A.; Sinchaipanid, N.; Faroongsarng, D. Spray-dried rice starch: Comparative evaluation of direct compression fillers. Drug Dev. Ind. Pharm. 1996, 22, 587–594. [Google Scholar] [CrossRef] [Scilit]
  41. Zhao, H.; Yu, Y.; Ni, N.; Zhao, L.; Lin, X.; Wang, Y.; Du, R.; Shen, L. A new parameter for characterization of tablet friability based on a systematical study of five excipients. Int. J. Pharm. 2022, 611, 121339. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  42. Conceição, A.P.; Sá, R.R.; da Silva, V.C.; Ferreira, M.S.; Cazedey, E.C.L.; Hemerson, I.F.; Magalhães, H.I.F.; Santos Júnior, A.F. A comparative study of propranolol release by in vitro dissolution profiles in pharmaceutical formulations. Dissolution Technol. 2018, 25, 54–61. [Google Scholar] [CrossRef] [Scilit]
Figure 1. SEM picture of native TS and TS agglomerates prepared using different contents of GTS.
Scipharm 94 00073 g001
Figure 2. Maximum force at 50% displacement of TS agglomerates prepared using different contents of GTS. Each value represents mean ± S.D., n = 10.
Scipharm 94 00073 g002
Figure 3. Heckel (a) and Kawakita (b) plots of TS agglomerates, and tensile strength (c) of TS agglomerate tablets using various % GTS as an agglomerating agent. Each point of Heckel and Kawakita plots is the mean from 3 tablets, and each point of the tensile strength is the mean ± S.D. of 3 tablets.
Scipharm 94 00073 g003
Figure 4. Surface morphology of tablet prepared from TS agglomerates with 2%GTS.
Scipharm 94 00073 g004
Figure 5. Carr’s index (a), angle of repose (b), maximum force at 50% displacement (c), and tablet hardness compressed at 12.3 MPa (d) of TS agglomerates prepared using PVP K30, SA and GTS in the concentration of 2% w/w. The data of PVP K30 and SA were obtained from references [12] and [20], respectively.
Scipharm 94 00073 g005
Figure 6. Drug dissolution profiles of PNL tablets prepared using TS agglomerates with different contents of GTS. Each point represents mean ± S.D., n = 3.
Scipharm 94 00073 g006
Figure 7. Hardness and friability of agglomerate tablets loaded with different contents of PNL in dilution potential testing. Each value of tablet hardness represents mean ± S.D., n = 6.
Scipharm 94 00073 g007
Figure 8. Drug dissolution profiles of agglomerate tablets loaded with different contents of PNL in dilution potential testing. Each point represents mean ± S.D., n = 3.
Scipharm 94 00073 g008
Table 1. Yield, density and particle flowability of TS agglomerates.
AgglomeratesYield (% w/w)Bulk Density a (g cm−3)Tapped Density a (g cm−3)True Density a (g cm−3)Carr’s Index a (%)Angle of Repose a (°)
0% GTS45.860.42 ± 0.010.49 ± 0.011.520 ± 0.00814.76 ± 2.0136.83 ± 0.74
1% GTS51.070.40 ± 0.010.46 ± 0.011.542 ± 0.00413.81 ± 1.2241.32 ± 0.17
2% GTS54.920.39 ± 0.010.45 ± 0.011.548 ± 0.01212.32 ± 1.1237.83 ± 1.00
3% GTS63.420.39 ± 0.010.44 ± 0.011.527 ± 0.01112.69 ± 0.9140.21 ± 0.81
5% GTS65.370.40 ± 0.010.46 ± 0.011.516 ± 0.01812.18 ± 1.6440.31 ± 0.53
a Data are mean ± S.D., n = 3.
Table 2. Heckel and Kawakita parameters of TS agglomerates.
TS AgglomeratesHeckel ParametersKawakita Parameters
K × 10−2 (1/MPa)APy (MPa)DaabPk (MPa)
0% GTS0.6430.924155.520.6030.6870.1865.376
1% GTS0.6540.842152.910.5690.6930.2084.808
2% GTS0.6660.799150.150.5500.6920.2284.386
3% GTS0.6830.770146.410.5370.6870.2324.310
5% GTS0.7010.823142.650.5610.6810.2354.255
Table 3. Characteristics of PNL-loaded tablets prepared using TS agglomerates.
AgglomeratesThickness a (mm)Hardness a (N)Disintegration Time b (min)T50% b (min)
0% GTS 2.62 ± 0.0111.93 ± 1.440.67 ± 0.019.94 ± 0.58
1% GTS2.62 ± 0.0117.33 ± 1.340.55 ± 0.064.56 ± 0.64
2% GTS2.73 ± 0.0321.74 ± 0.740.28 ± 0.011.34 ± 0.07
3% GTS2.75 ± 0.0331.54 ± 3.310.24 ± 0.011.72 ± 0.11
5% GTS2.70 ± 0.0227.95 ± 2.540.23 ± 0.061.32 ± 0.09
a Data are mean ± S.D., n = 6; b Data are mean ± S.D., n = 3.
Table 4. Physical properties of tablets prepared using TS agglomerate with 3% GTS as direct compression filler.
PNL Added (% w/w)Thickness a
(mm)
Tablet Weight b (mg)PNL Content in Tablet c (% w/w)Disintegration Time c (min)T50% c (min)
03.59 ± 0.02355.91 ± 0.57
(%RSD = 0.16)
-3.27 ± 0.16-
103.49 ± 0.03351.02 ± 1.06
(%RSD = 0.30)
100.14 ± 0.430.96 ± 0.012.90 ± 0.16
203.51 ± 0.05348.35 ± 3.60
(%RSD = 1.03)
97.16 ± 2.320.99 ± 0.022.33 ± 0.32
303.51 ± 0.02348.93 ± 1.89
(%RSD = 0.54)
96.60 ± 0.540.43 ± 0.101.60 ± 0.26
403.54 ± 0.02351.02 ± 3.33
(%RSD = 0.95)
101.85 ± 2.580.51 ± 0.071.60 ± 0.29
a Data are mean ± SD., n = 6; b Data are mean ± SD., n = 20; c Data are mean ± SD., n = 3; RSD = relative standard deviation.
Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content.