


Abstract
The escalation of thermal runaway in lithium-ion batteries presents severe safety hazards that necessitate advanced monitoring protocols to ensure early warning of potential failures. Carbon dioxide (CO2) is released during preliminary decomposition well before catastrophic failure occurs, thereby providing a strategic advantage for early-stage warning. Consequently, identifying materials with high-selective CO2 recognition is an essential prerequisite for developing reliable sensing platforms. This study integrates Grand Canonical Monte Carlo simulations with Random Forest (RF) models to systematically screen 1470 MOFs from the CoRE-MOF 2019 database. The screening process evaluates selective CO2 recognition under multicomponent competitive adsorption conditions involving CO2, C2H4, and O2. The performance evaluation is based on working capacity, selectivity, and the trade-off between working capacity and selectivity (TSN). The RF model achieves high predictive accuracy, with tested R2 exceeding 0.92 on the test samples. Shapley Additive Explanations (SHAP) interpretability analysis identifies Q0st(CO2), Q0st(C2H4), WEPA, KH(C2H4), and ETR as key performance drivers. The results indicate that CO2 selectivity is constrained by the binding strength of competing C2H4. Optimal materials tend to have hard Lewis acid centers and polar inorganic clusters to minimize non-specific π-interactions with interfering species. Top-performing MOFs require balanced structural features, concentrating in moderate surface areas (965–1975 m2/g), narrow pore windows (PLD ≈ 4–7 Å, LCD ≈ 5.5–9.6 Å), high void fractions above 0.6, and low densities below 1.3 g/cm3. AJOTEY emerges as the optimal candidate with a TSN of 6.43 mol/kg, combining substantial working capacity (4.57 mol/kg) with strong selectivity (25.52). These results will accelerate the discovery of sensing materials and provide a practical pathway for MOF-based CO2 sensor development to enhance lithium-ion battery safety.
1. Introduction
Lithium-ion batteries are central to the global shift towards electric mobility and stationary energy storage systems [1,2]. However, thermal runaway remains a critical safety concern that can trigger catastrophic failures including fires and explosions [3,4]. The thermal runaway process unfolds in several stages, each associated with distinct gas emissions [5]. Carbon dioxide (CO2) is the primary gas released during the early stages, emerging at temperatures as low as 80–120 °C when the solid electrolyte interphase starts to break down. This initial CO2 emission offers a critical opportunity for intervention before the temperature rises further [6]. In contrast, hydrogen (H2) is released in large quantities above 200 °C. This occurs in the later stages, when the electrolyte decomposes violently. Most existing safety monitoring strategies therefore rely on H2 detection, by which point the battery has already entered a highly dangerous state [7,8]. By the time H2 reaches detectable levels, thermal runaway has already progressed significantly, making intervention extremely difficult. Recent studies have shown that CO2-selective systems can provide warning signals several minutes earlier than H2-based approaches [9]. This early warning provides time for cooling, power disconnection, or evacuation [10]. Taken together, these results highlight CO2 recognition as a more effective route for early-stage thermal runaway warning compared with conventional H2-based strategies. This effectiveness places the primary emphasis on developing materials with intrinsic and selective CO2 recognition capability.
Metal-organic frameworks (MOFs) are porous crystalline materials with high specific surface areas and tunable pore structures, making them promising candidates for CO2 sensing in battery safety applications [11]. When employed as gas-responsive materials, their performance is governed by sensitivity, selectivity, response kinetics, and long-term stability. These properties can be systematically tuned through rational structural design. For example, HKUST-1 utilizes open Cu2+ sites to enable strong chemisorption [12,13], while NH2-Mg-MOF-74 employs ethylenediamine groups to amplify capacitive response [14]. Due to the complex gas mixtures released by batteries, selectivity is crucial for early thermal runaway detection, as CO2 must be distinguished from interfering species like C2H4 and O2. Some MOFs have demonstrated promising selectivity through molecular sieving effects. ZIF-8 with its narrow 3.4 Å pores selectively admits CO2 while excluding larger molecules. MUF-16 shows inverse selectivity over C2H4 with uptake ratios reaching 15.6 [7,10,15]. Despite these successes, identifying the most suitable MOF candidates from the vast structural landscape remains a formidable challenge.
Addressing this challenge increasingly relies on data-driven strategies that enable efficient exploration of large materials spaces. Currently, data-driven methodologies are revolutionizing battery science. Advanced deep learning techniques have been extensively applied to enhance battery thermal management and realize early safety warnings [16], while emerging large language models are accelerating material discovery and experimental design by mining vast databases [17]. Over 100,000 MOF structures have been synthesized to date, with millions more theoretically possible through different metal-ligand combinations [18,19,20,21,22]. Experimentally testing each candidate for CO2 selectivity against interfering gases like C2H4 and O2 would be prohibitively time-consuming and expensive [23,24,25]. Traditional computational screening relies on high-throughput molecular simulations such as Grand Canonical Monte Carlo (GCMC) to calculate gas adsorption properties [26]. While GCMC provides accurate predictions, comprehensive evaluation of large-scale databases like CoRE-MOF with over 14,000 structures remains computationally prohibitive [27,28,29]. Machine learning (ML) offers a transformative solution by learning structure-property relationships from limited simulation data [30]. Models trained on structural descriptors such as pore size, surface area, and functional group distribution can rapidly predict adsorption selectivity for thousands of MOF candidates [31,32]. This computational strategy enables efficient identification of MOFs optimized for detecting the early CO2 signature of thermal runaway in battery systems.
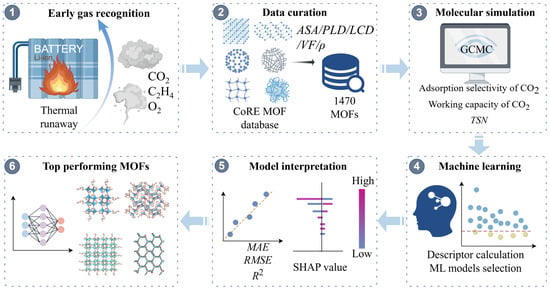
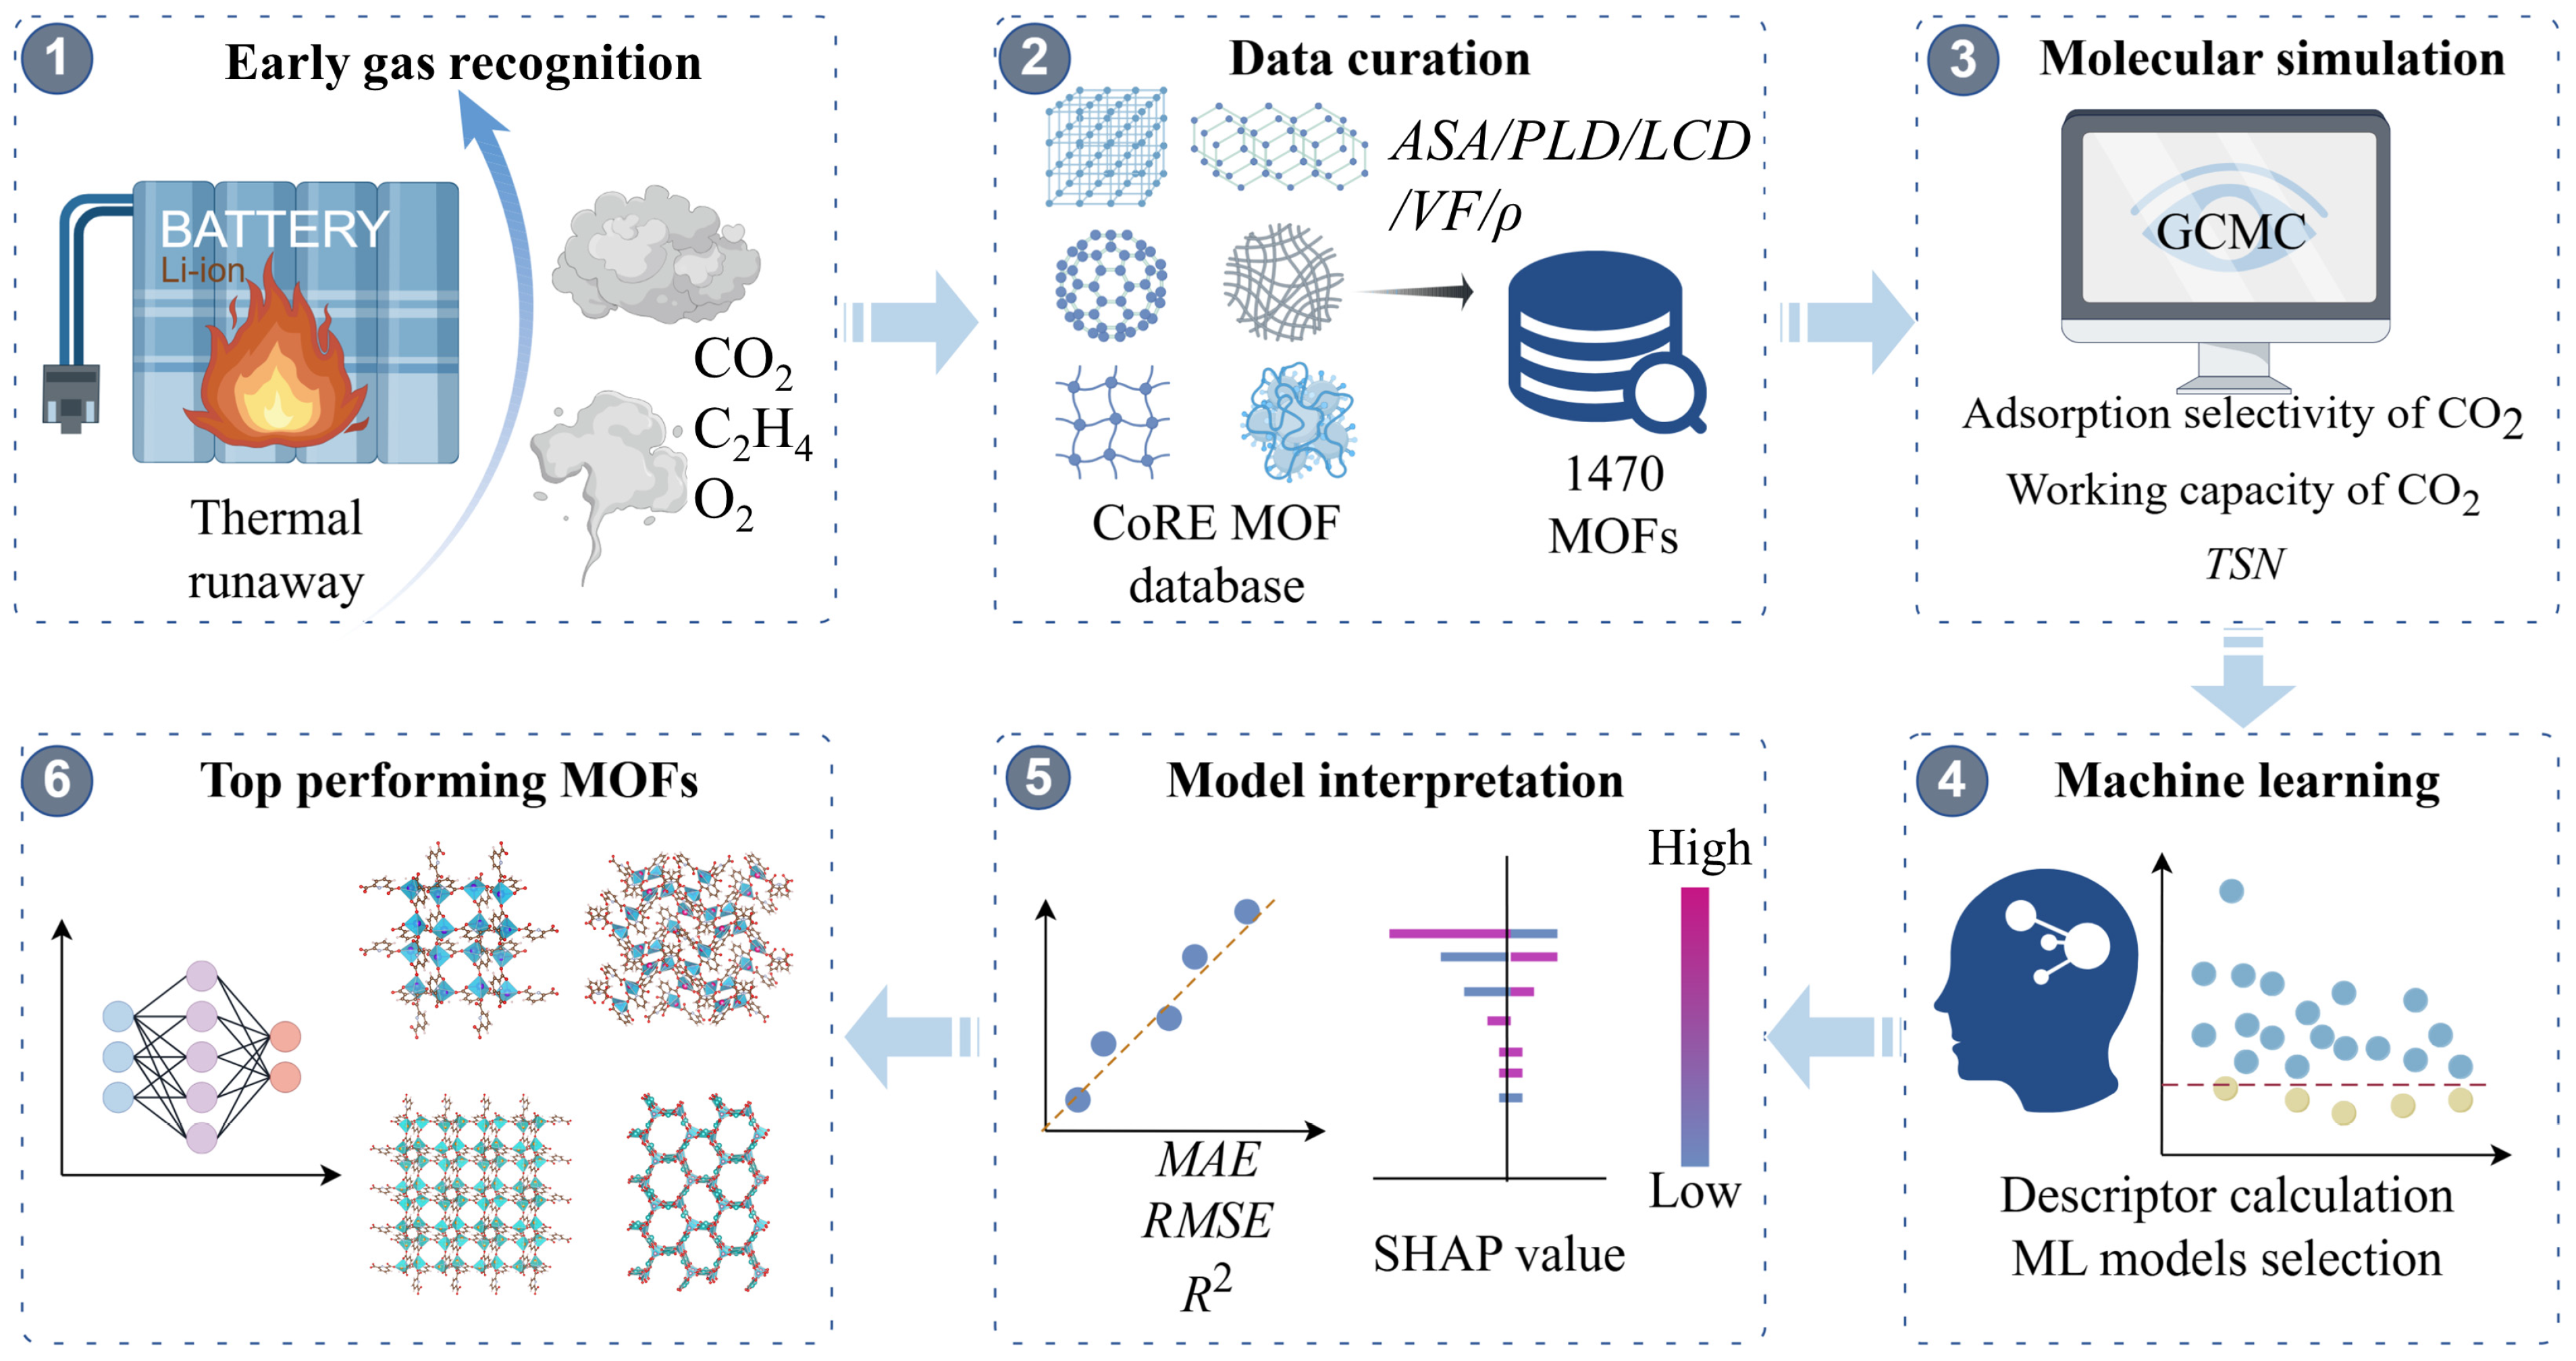
In this study, GCMC simulations combined with Random Forest (RF) models were used to systematically screen 1470 MOF materials for CO2 recognition in battery thermal runaway scenarios (Scheme 1). We assess working capacity, selectivity, and overall performance under multicomponent competitive adsorption conditions, which simulate the complex gas mixtures released during early battery failure. Analysis of key structural descriptors reveals non-linear relationships between MOF architecture and CO2 adsorption performance. Model interpretability techniques identify the dominant physicochemical factors governing CO2 selectivity against interfering gases. High-performance MOF candidates for selective CO2 recognition were identified. Such capabilities are crucial in the early stages of lithium-ion battery thermal runaway, offering a new pathway for developing advanced battery safety systems.
Scheme 1.
Computational workflow for ML-guided screening of MOFs for CO2 recognition in battery thermal runaway applications by figdraw.com.
2. Computational Details
2.1. Database and Screening Process for MOFs
Structural and chemical descriptors were calculated for 1470 MOFs as listed in Table S1. The MOF dataset was initially selected from the CORE-MOF 2019 database totaling 6603 MOFs. To characterize the geometric and size attributes of the internal pores in MOFs, five structural descriptors were computed like largest cavity diameter (LCD), pore limiting diameter (PLD), accessible surface area (ASA), density (ρ), and void fraction (VF). These descriptors were calculated using the Zeo++0.3 software, with He and N2 as probes (kinetic radius: 1.32 Å and 1.86 Å, respectively) to estimate VF and ASA.
In addition to these structural descriptors, adsorption enthalpy and Henry’s coefficients (KH) under infinite dilution conditions were incorporated as chemical descriptors of the MOFs. Furthermore, various types of atomic counts were extracted from the crystal structures of the MOFs, from which additional chemical descriptors were derived. These descriptors include the type and number of each atom, total unsaturation degree, metal percentage, oxygen-to-metal ratio, electronegativity ratio, weighted electronegativity per atom, and nitrogen-to-oxygen ratio.
The screening of the MOFs was based on the following criteria: (1) PLD > 3.3 Å, (2) 950 m2/g < ASA < 5000 m2/g, (3) 0.1 < VF < 0.7, and (4) KH(H2O) < 2.6·10−6 mol/(kg·Pa). This process resulted in the selection of 1470 MOFs for further analysis.
2.2. Simulation Details for GCMC
In the initial stage of thermal runaway in lithium-ion batteries, the temperature continuously rises due to the decomposition of the solid electrolyte interphase (SEI) between 80–120 °C, leading to the first reaction. When the temperature approaches 90 °C, the decomposition of SEI begins to release a significant amount of heat and gases (CO2, C2H4, and O2) [33]. The reaction is as follows:
(CH2OCO2Li)2 → Li2CO3 + C2H4↑ + CO2↑ + 1/2O2↑
The initial stage of thermal runaway is simulated as a CO2/C2H4/O2 mixed gas with mole fractions of 0.4/0.4/0.2. Separation is performed using the vacuum swing adsorption (VSA) process, with adsorption at 1 bar and desorption at 0.1 bar. The operating temperature is 363 K (approximately 90 °C).
To describe the adsorbate-adsorbate interactions, the universal force field was applied to the MOFs [34]. The interactions between adsorbates were described using the Lorentz-Berthelot mixing rules. In the calculations for each structure, the supercell of the MOF was used, ensuring that the simulation box had dimensions at least twice the cutoff radius along each crystallographic direction. Non-bonded interactions were represented by the Lennard-Jones potential with a cutoff radius of 12.8 Å, while Coulombic interactions were computed using the Ewald summation method, with a relative error of 106. The Monte Carlo moves in the GCMC simulation included translation, rotation, addition/deletion, re-insertion, and identity change, all attempted with equal probability. The initialization run was set to a total of 1 × 10⁴ cycles, followed by 3 × 10⁴ cycles for the production run.
2.3. Calculation of Chemical Descriptors by RASPA
All simulations assume rigid MOF structures and equilibrium adsorption under idealized dry gas conditions, which allows internally consistent comparison across chemically diverse MOF datasets. Framework flexibility, humidity, electrolyte vapors, and time-dependent response will be addressed during device-level evaluation rather than at the material screening stage. The same assumptions are applied uniformly to all candidate materials, ensuring that the identified trends and structure-property relationships for CO2 recognition are robust within the defined operating conditions.
Adsorption-related chemical descriptors were calculated using the RASPA2.0 software [35]. These descriptors include Henry’s law coefficients (KH) and adsorption enthalpies at infinite dilution (Q0st) for CO2, C2H4, and O2, which collectively describe adsorption affinity and energetic interactions under low-pressure conditions. Henry’s law coefficient KH reflects the initial adsorption affinity and governs competitive uptake behavior in the low-pressure regime relevant to early thermal runaway conditions. In contrast, Q0st characterizes the intrinsic interaction strength between gas molecules and the framework and provides thermodynamic insight into adsorption selectivity. KH values were obtained using the Widom test particle insertion method (with a Widom probability set to 1) with 10,000 Monte Carlo cycles, while Q0st values were calculated from additional simulations of 20,000 cycles under 367 K [36].
To account for moisture effects that may arise in realistic battery environments, hydrophobicity was introduced as an explicit screening constraint. KH(H2O) was calculated for all 1470 candidate MOFs at 363 K using the same Widom insertion protocol. Hydrophobic materials were identified using a threshold of KH(H2O) < 2.6 × 10⁻⁶ mol/(kg·Pa), as established in previous studies [23]. Applying this criterion effectively excludes hydrophilic structures and ensures that the selected candidates possess intrinsic resistance to water interference. Supporting Information (SI) and Table S2 provide other relevant chemical descriptors.
2.4. Adsorption Performance Metrics
Three evaluation metrics were employed: the adsorption selectivity of CO2 over C2H4 and O2 (S), the working capacity of CO2 (ΔN), and the trade-off between working capacity and selectivity (TSN) [23]. The specific equations are presented in Table 1, where and represent component in the adsorption and gas phase, respectively. and are the CO2 loading under the adsorption and desorption pressures, respectively.
Table 1.
Metrics used to evaluate the MOF adsorbents.
3. Result and Discussion
3.1. Distributions of Structural Descriptors
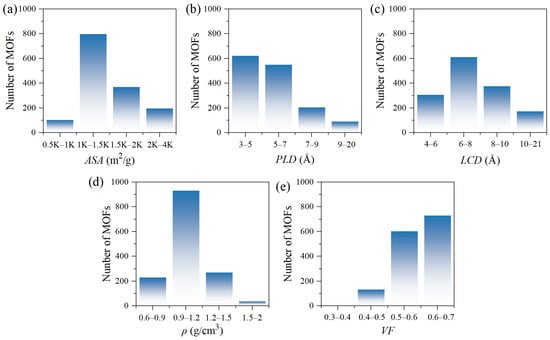
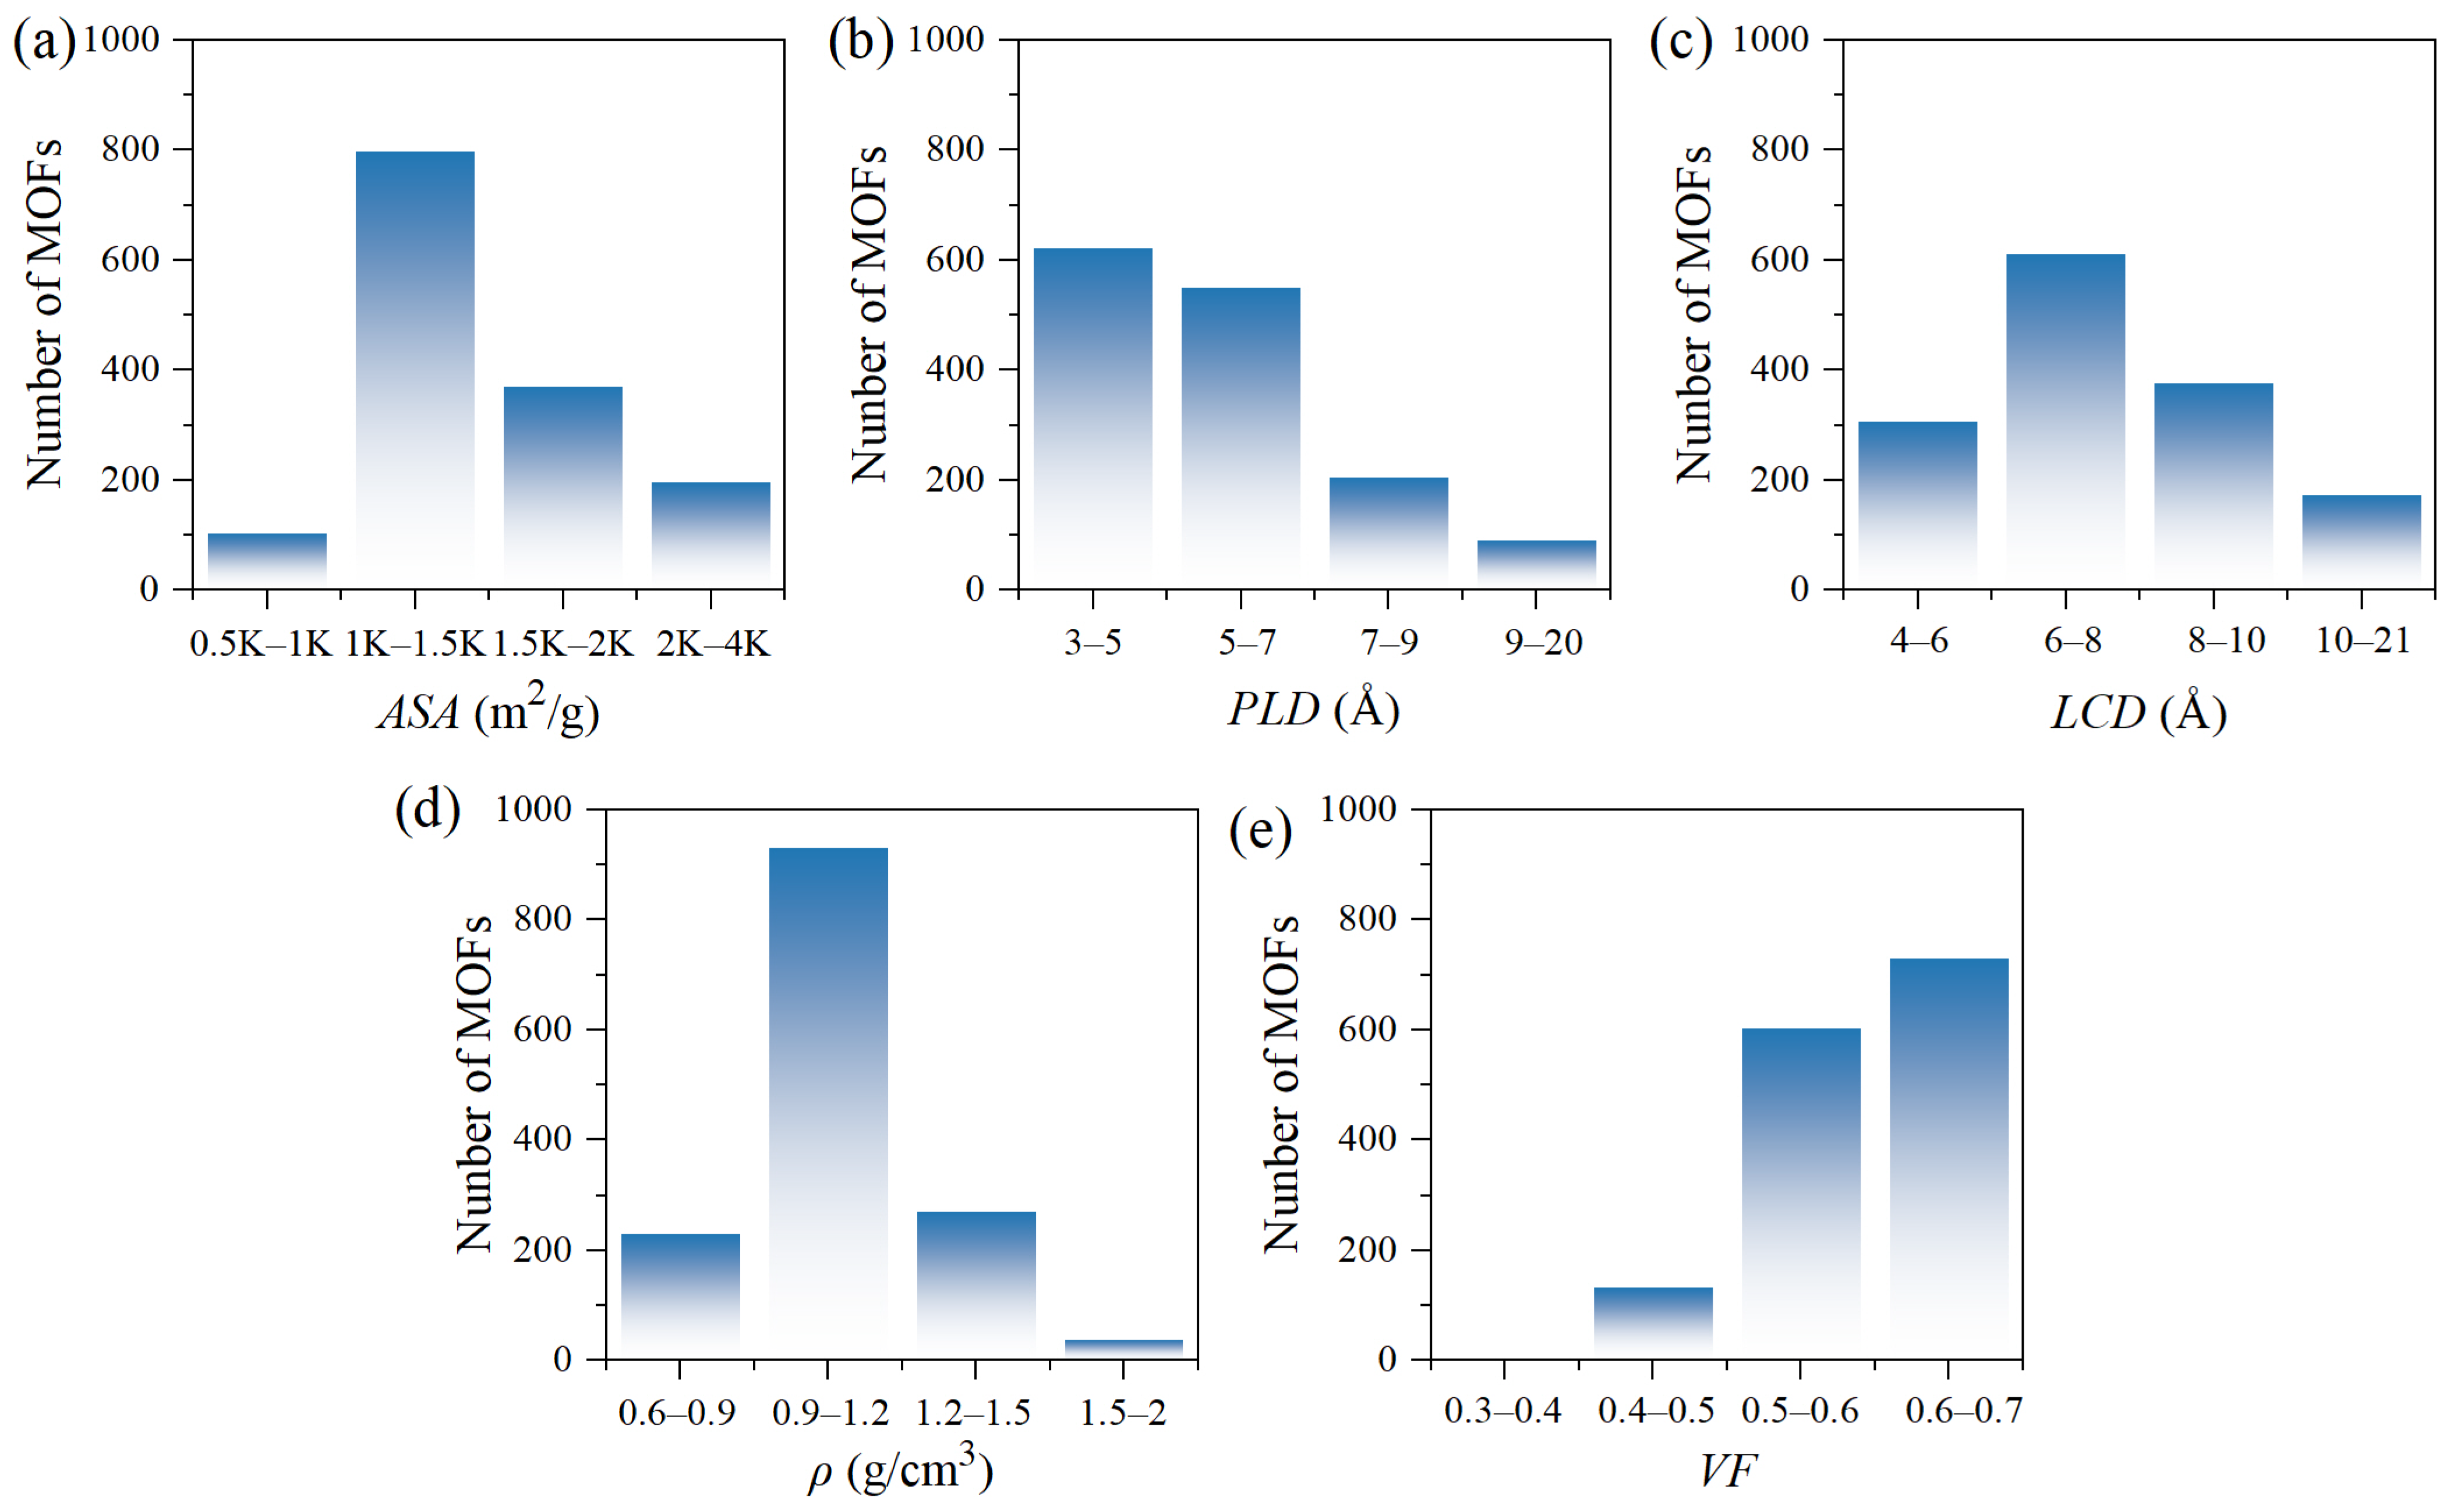
The structural diversity of the 1470 MOFs in the dataset plays a critical role in ensuring robust ML models with strong generalization capability. Figure 1 presents histograms of five key structural descriptors. These distributions cover broad value ranges and align well with the requirements for selective CO2 adsorption under low-pressure competitive conditions. ASA is a key factor influencing physisorption capacity, with values ranging from around 500 to over 4000 m2/g. Most materials cluster between 1000 and 2000 m2/g as shown in Figure 1a. This wide distribution enables the model to capture the complex relationship between surface area and performance. This challenges the oversimplified view that higher ASA always leads to higher loading [37]. MOFs with PLD in the 3–7 Å range are the most abundant (Figure 1b), while those with LCD show a broader distribution extending up to approximately 20 Å (Figure 1c). The target gases have kinetic diameters of 3.3 Å for CO2, 3.9 Å for C2H4, and 3.64 Å for O2. Most MOFs therefore possess pore dimensions closely matching these molecular sizes, providing the structural basis for molecular sieving and diffusion selectivity. Pores significantly smaller than these dimensions restrict gas entry, while excessively large pores diminish selectivity [38]. The wide PLD and LCD distributions allow the model to learn structure-performance relationships across various architectures, rather than being limited to a narrow structural range. The highest number of MOFs falls within the ρ range of 0.9 to 1.2 g/cm3 (Figure 1d) indicating greater framework compactness, which typically correlates inversely with porosity. VF remains predominantly high, with most materials between 0.5 and 0.7 (Figure 1e), indicating substantial internal space for gas adsorption. These descriptors collectively capture the structural features governing CO2 selectivity in competitive adsorption scenarios.
Figure 1.
Histograms showing the distribution of key structural descriptors for 1470 MOFs: (a) ASA, (b) PLD, (c) LCD, (d) ρ, and (e) VF.
Key descriptors span a wide range of values, which enables ML models to capture diverse structure-performance relationships across various MOF architectures, rather than being restricted to specific structural types. The pore size distributions are primarily concentrated in the microporous range, matching the dimensions of target gases like CO2 and C2H4. Along with favorable pore volume and surface area distributions, these features highlight the database’s inclusion of numerous materials with strong adsorption potential. This structural diversity not only enhances model generalization but also supports the rational and application-oriented nature of the data selection process.
3.2. Descriptor-Dependent Adsorption Trends in Low-Pressure Multicomponent CO2 Recognition
Structure-performance relationships reveal non-intuitive trends for low-pressure multicomponent CO2 recognition (Figure S1). Working capacity does not show a simple positive correlation with ASA, ρ, or VF. Instead, top performers are found at moderate ASA values between 950 and 1750 m2/g. They also fall within optimal pore ranges of PLD (4–8 Å) and LCD (5–9 Å), balancing diffusion access with strong van der Waals interactions (Figure S1). Selectivity favors smaller pores and lower ASA to enhance molecular confinement and restrict interfering gases (Figure S1). The composite metric TSN integrates both capacity and selectivity, which exhibits the sharpest structural dependencies. High-performing MOFs occupy narrow pore size ranges of 4–7 Å PLD and 5.5–9.6 Å LCD, combined with moderate ASA, high VF above 0.6, and low ρ below 1.3 g/cm3 (Figure S1).
These findings differ markedly from traditional high-pressure single-component adsorption studies which uptake scales directly with surface area. In the targeted low-partial-pressure competitive environment, efficient site utilization and precise pore tuning outweigh any single descriptor. ML uncovers these nuanced structure-performance relationships that conventional heuristic screening would miss. This enables the identification of MOF candidates optimized for selective CO2 recognition in early lithium-ion battery thermal runaway warning systems.
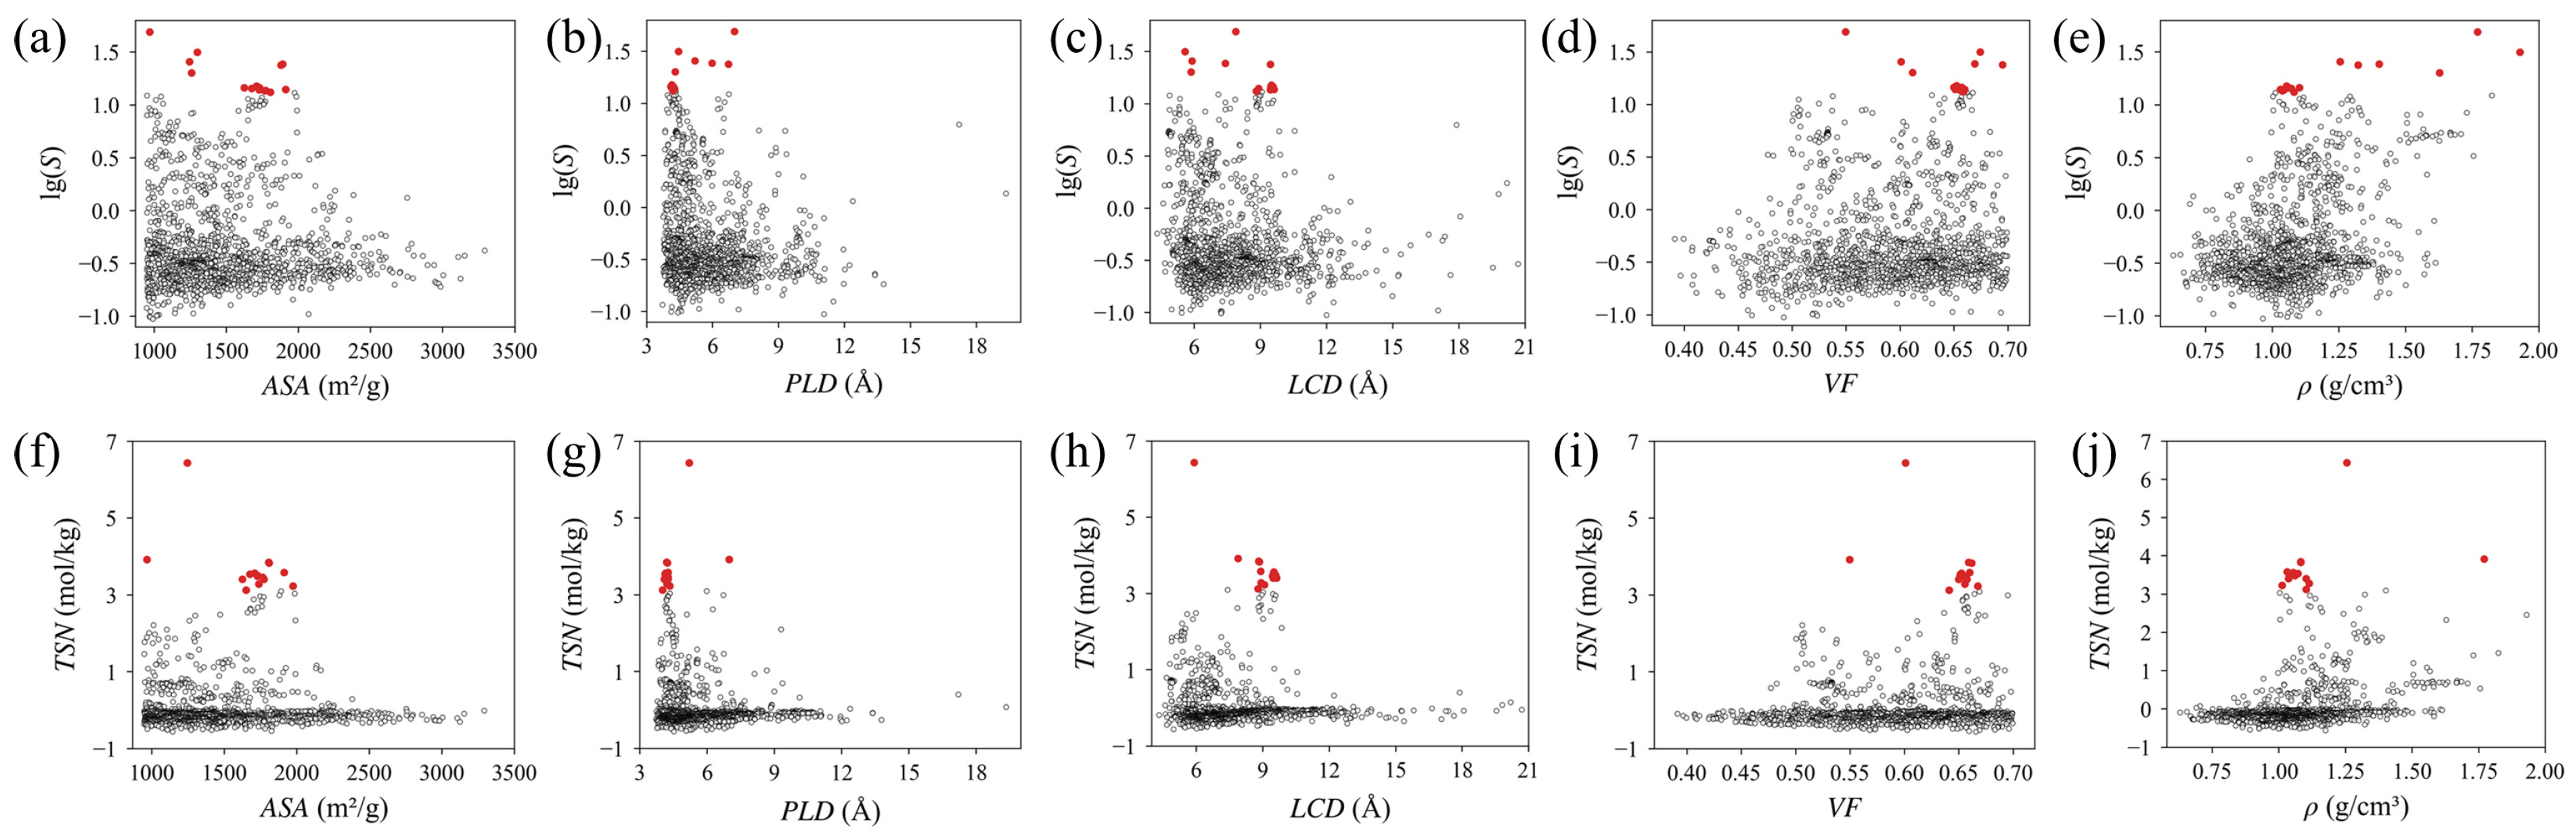
Selectivity exhibits distinct structural dependencies favoring confined environments (Figure 2). Although selectivity generally increases as ASA decreases, the highest-performing MOFs cluster at moderate values rather than at the minimum extremes (Figure 2a). Excessively open structures reduce confinement effects that enhance CO2 binding relative to C2H4 and O2. Furthermore, pore dimensions strongly influence selectivity, with performance decreasing as pore sizes increase (Figure 2b,c). Top-performing MOFs cluster within a moderately small pore range that provides sufficient confinement to strengthen CO2 interactions without restricting diffusion. In contrast, VF and ρ show less pronounced correlations with selectivity (Figure 2d,e), though high performers still favor moderately high VF above 0.5 and low ρ below 2 g/cm3. These trends indicate that superior CO2 selectivity under competitive low-pressure conditions emerges from balanced pore confinement that optimizes interaction strength differences among competing gases. Consequently, such selectivity is governed by these discrete local environments rather than the broad maximization of surface area or pore volume.
Figure 2.
Scatter plots showing the relationship between CO2 selectivity (lg(S)) and five key structural descriptors: (a) ASA, (b) PLD, (c) LCD, (d) VF, (e) ρ. The relationship between TSN and five key structural descriptors: (f) ASA, (g) PLD, (h) LCD, (i) VF, (j) ρ. The top 15 MOFs are highlighted as red points, and the remaining MOFs are shown in gray points.
TSN shows clear structural dependencies for selective CO2 recognition under thermal runaway conditions. High-performing MOFs concentrate in the moderate ASA range of approximately 965 to 1975 m2/g (Figure 2f). This range balances CO2 working capacity with selectivity against C2H4 and O2. While increasing ASA provides more total adsorption sites, it simultaneously compromises selectivity by facilitating indiscriminate gas uptake. Pore size descriptors PLD and LCD exhibit sharply peaked relationships with TSN, revealing a narrow optimal window (Figure 2g,h). High-TSN materials cluster almost exclusively within PLD ≈ 4–7 Å and LCD ≈ 5.5–9.6 Å. Within these specific dimensions, pore confinement effects preferentially strengthen CO2 adsorption relative to the slightly larger C2H4 (3.9 Å) and O2 (3.64 Å). This behavior confirms that selectivity originates from enhanced interaction strengths within confined spaces rather than simplistic size exclusion. Conversely, TSN rapidly declines when pore dimensions exceed PLD > 7 Å or LCD > 10 Å, as enlarged channels permit competing gases to adsorb with similar affinities. TSN also shows clear preferences for VF and ρ (Figure 2i,j). High-performing MOFs favor high VF above 0.6 and low framework densities below 1.3 g/cm3. Such a combination ensures maximum site exposure and mass transfer efficiency without sacrificing gravimetric capacity. Collectively, these trends establish a precise profile for MOF design: moderate surface area, 4–7 Å pore windows, and high-porosity, low-density frameworks. This structural profile holds promise for providing explicit guidance in developing CO2 sensors for battery safety applications.
3.3. Predictive Performance of the RF Model
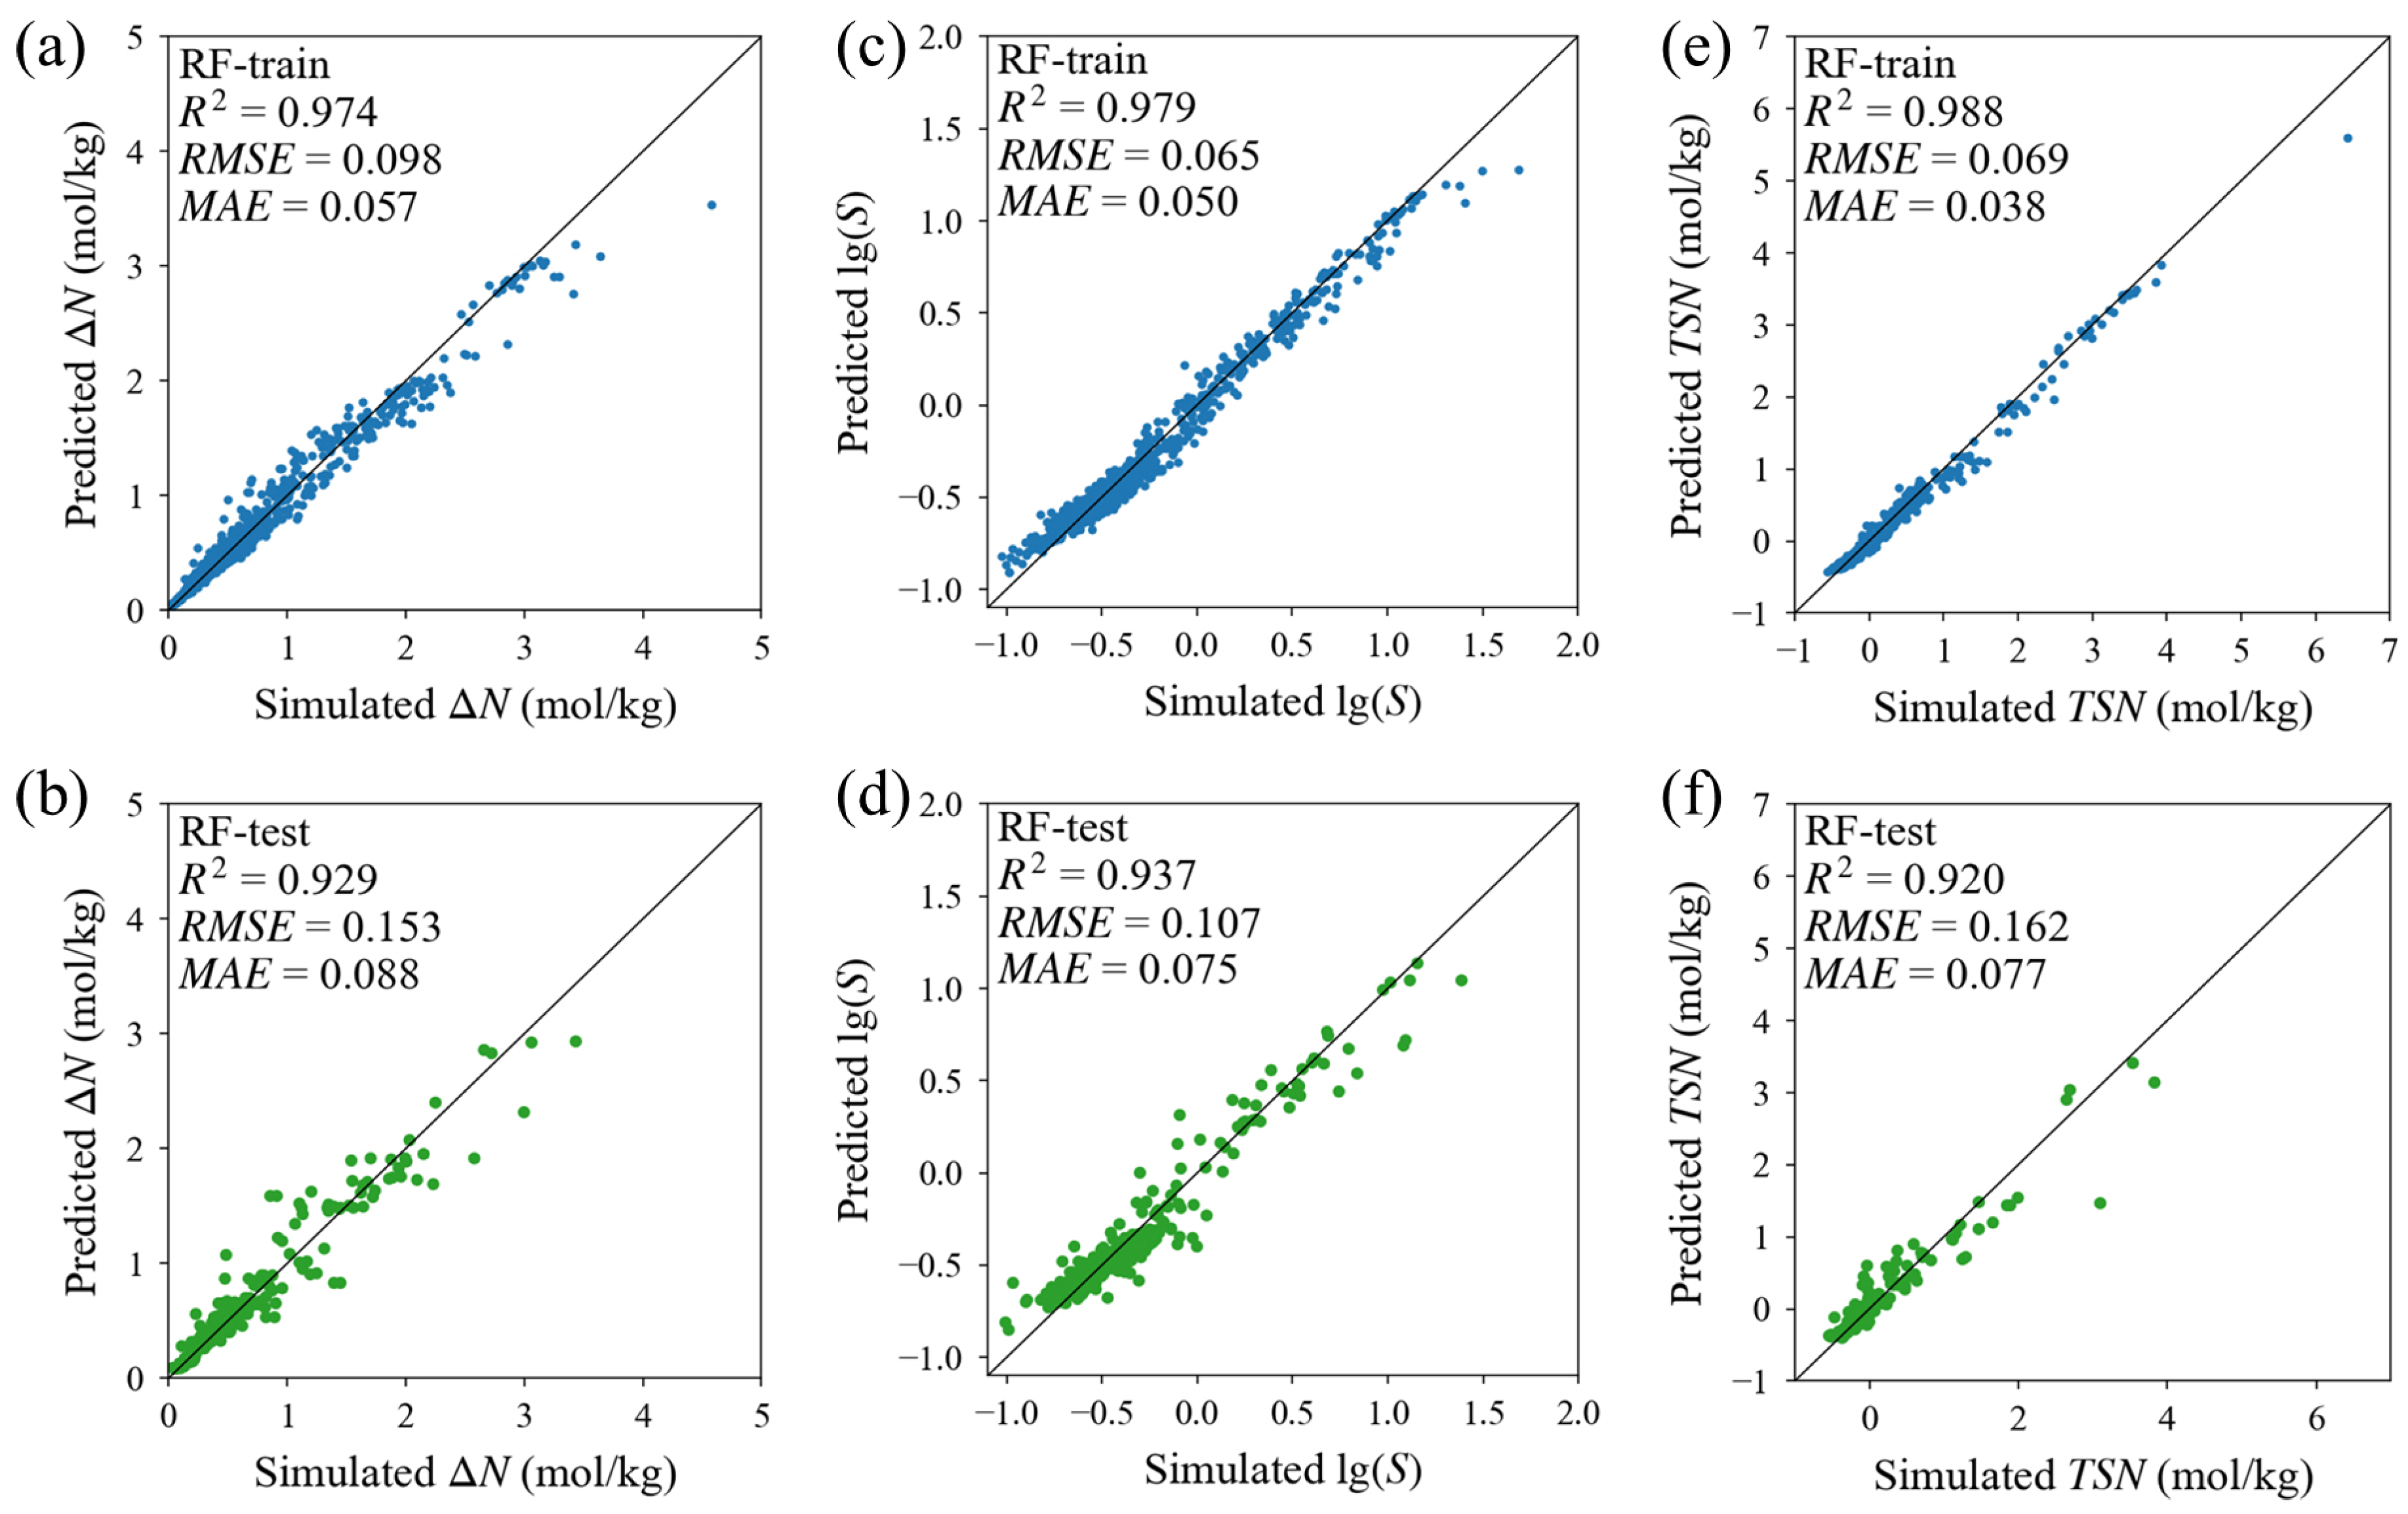
This study systematically compared three machine learning algorithms, including Decision Tree (DT), Random Forest (RF), and Extreme Gradient Boosting (XGBoost), to identify a robust predictive model for CO2/C2H4/O2 separation performance. The dataset was randomly divided into a training set comprising 80% of the data and an independent testing set comprising the remaining 20%. Bayesian hyperparameter optimization combined with 10-fold cross validation was applied during model training to improve robustness and reduce the risk of overfitting. The predictive performance of the optimized models on the independent testing set is summarized in Table S3. Although XGBoost achieved extremely high fitting accuracy on the training set with R2 values exceeding 0.99, its performance declined markedly on the testing set, with a testing R2 of approximately 0.82 for lg(S). This behavior indicates limited generalization capability. In contrast, the RF model demonstrated superior generalizability and consistency across all individual metrics. For instance, RF model maintained high and stable testing R2 values for ΔN (0.929), lg(S) (0.937), and TSN (0.920), consistently outperforming or matching other models in the testing phase. Based on this robust balance between predictive accuracy and generalizability, the RF model was selected for subsequent analysis.
Detailed evaluation of the RF model (Figure 3) confirms its high predictive efficacy. For working capacity ΔN, the narrow gap between training and testing metrics signifies the model’s ability to internalize intrinsic structure-property relationships rather than merely memorizing data (Figure 3a,b). Predictive accuracy is further enhanced for lg(S) (R2 = 0.937), where the logarithmic transformation successfully normalized the distribution to facilitate the precise identification of highly selective MOFs (Figure 3d). Finally, the composite TSN metric exhibits strong performance (R2 = 0.920) with data points tightly clustered around the diagonal (Figure 3e,f). This high accuracy across all derived metrics validates the credibility of the top-ranked candidates identified from the database (Tables S4 and S5).
Figure 3.
Comparison between predicted and simulated values of (a,b) ΔN, (c,d) lg(S), and (e,f) TSN on the training and testing sets using the RF model.
The strong predictive consistency of the RF model across all three metrics confirms that TSN can be reliably used as a screening indicator linking adsorption capacity and selectivity. This level of predictive accuracy provides a data-driven basis for identifying MOFs that simultaneously exhibit high CO2 working capacity and strong competitive adsorption behavior.
3.4. Interpretation of Structure-Property Relationships Using SHAP Analysis
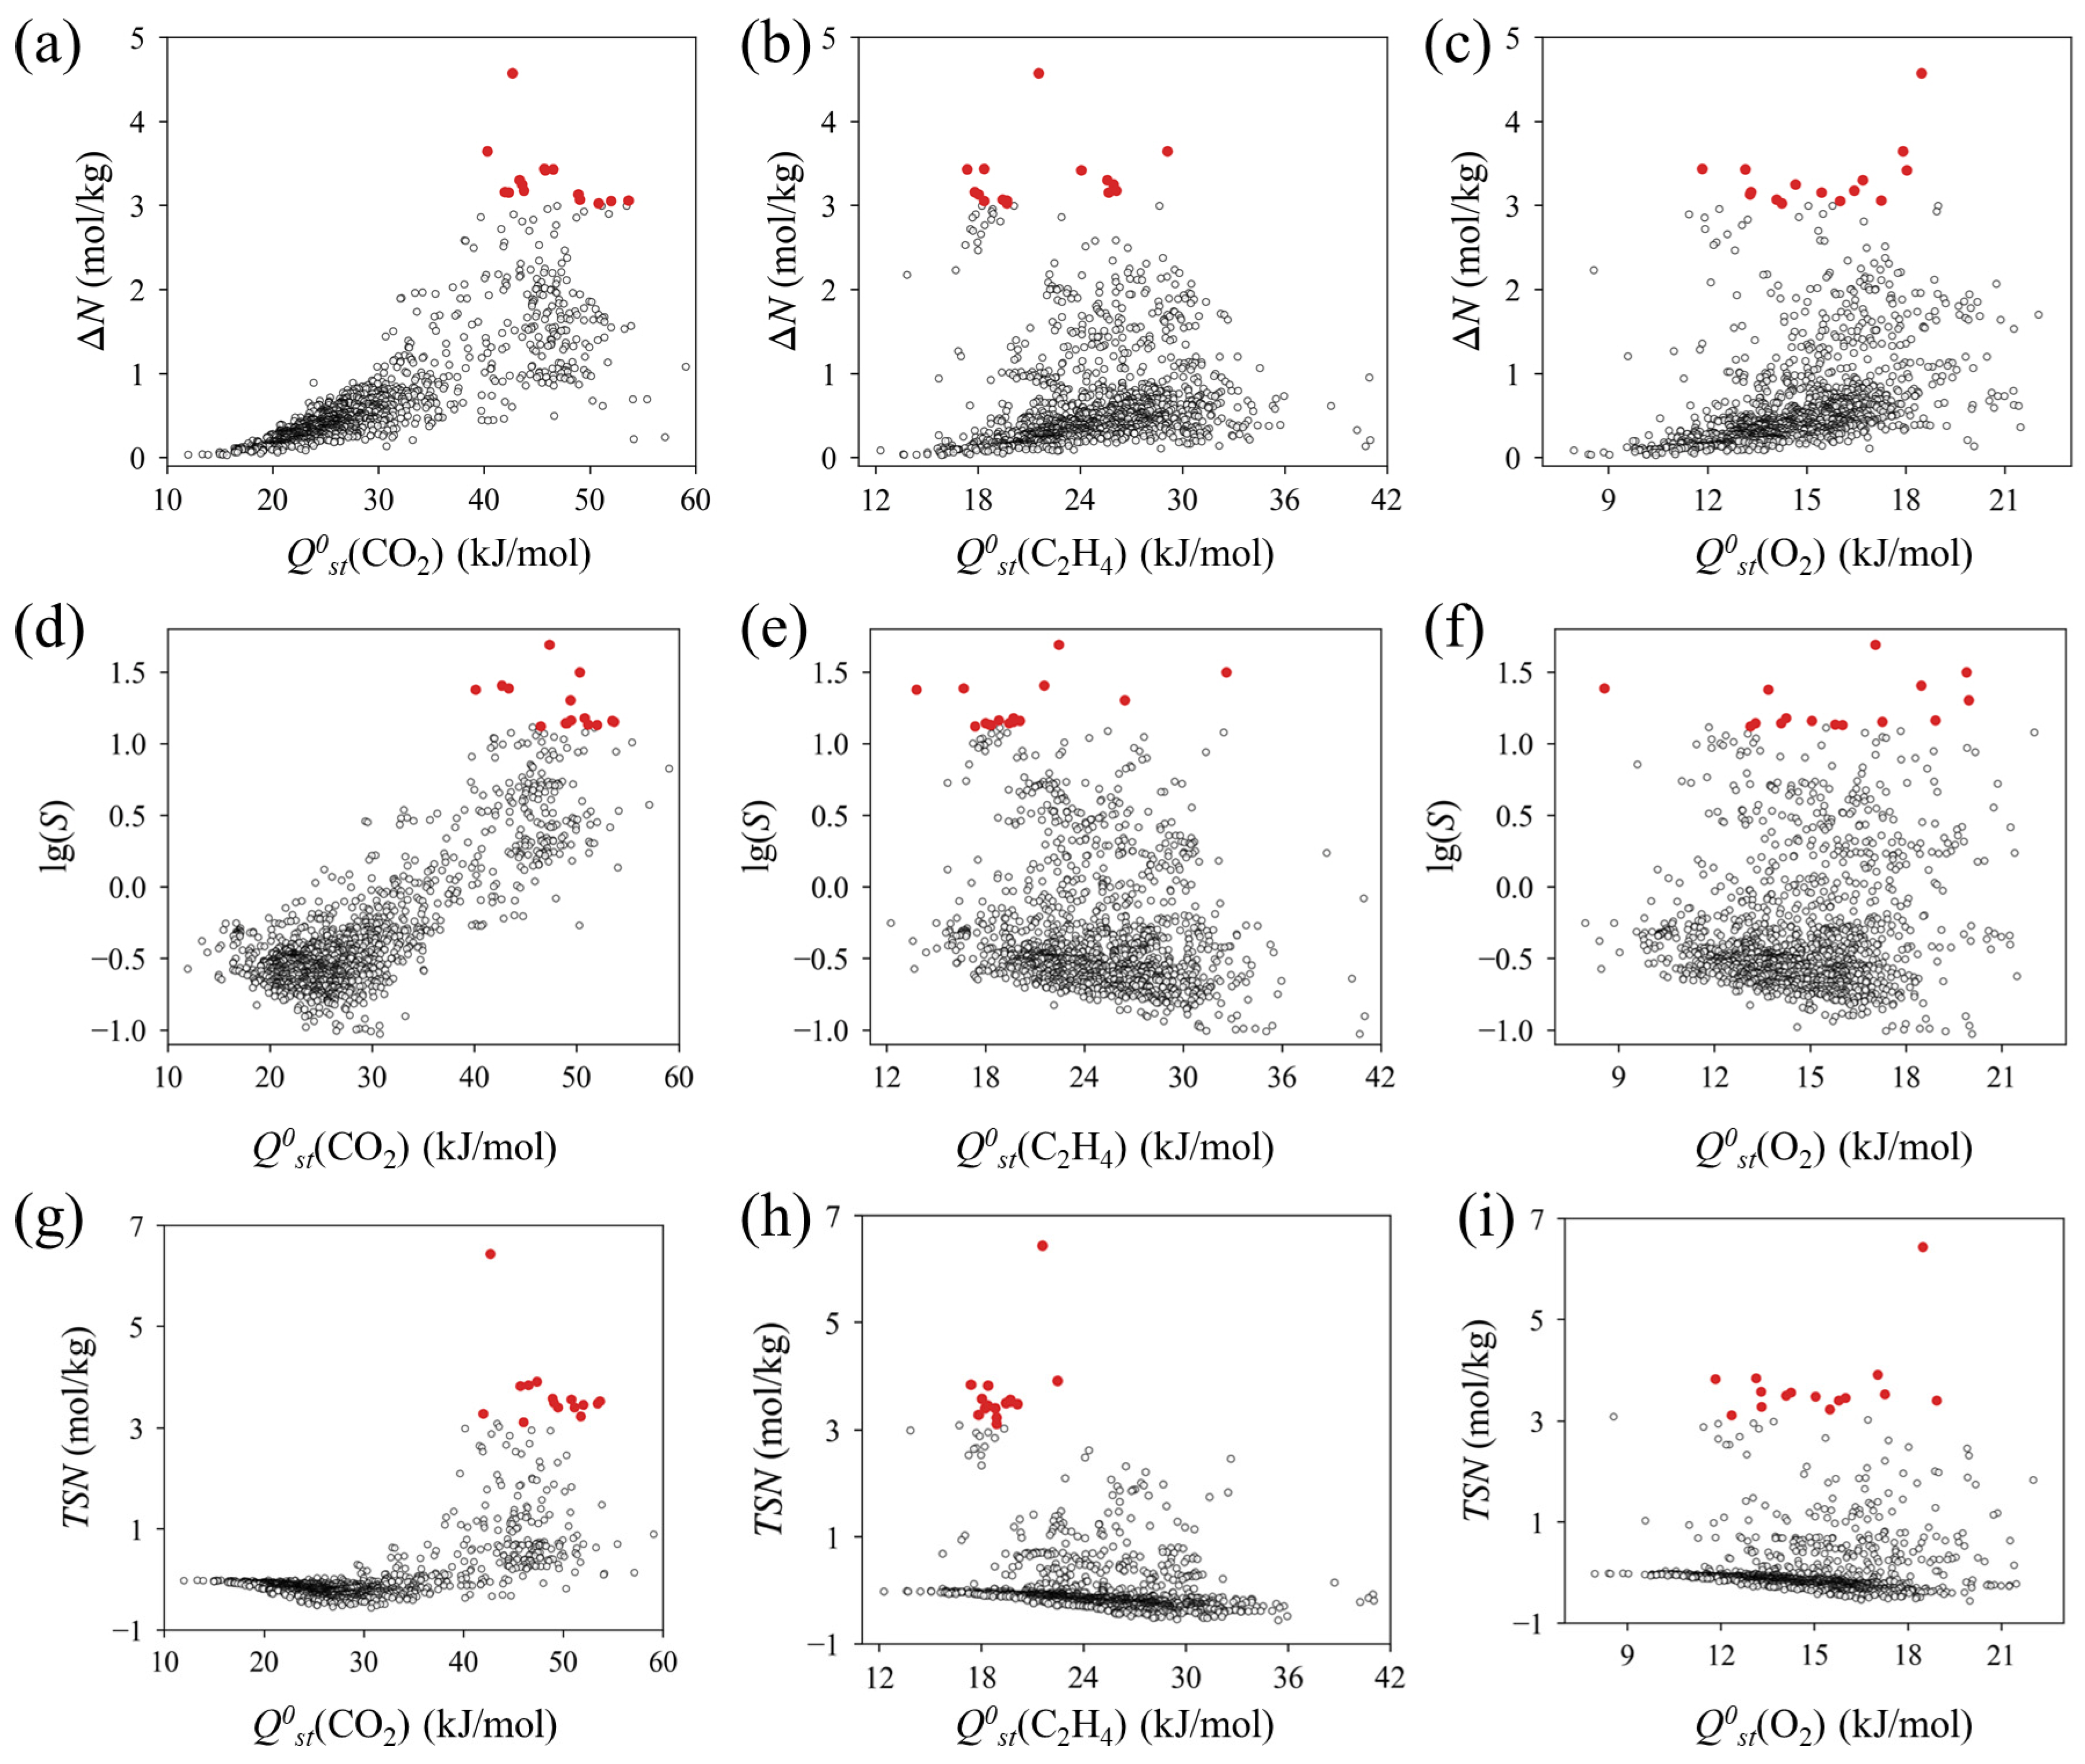
Prior to the multivariate SHAP analysis, we examined the univariate correlations to establish the fundamental physical bounds governing separation performance. As shown in Figure 4a–c, ΔN displays a non-monotonic, volcano-like trend relative to Q0st. This behavior aligns with the Sabatier principle, reflecting a critical balance that interactions must be strong enough to drive uptake but moderate enough to facilitate efficient regeneration [39]. For CO2, the highest capacities occur within an optimal window (~42.7 kJ/mol) instead of the theoretical maximum (Figure 4a). Similarly, C2H4 capacity peaks at an intermediate Q0st (C2H4) of 21.6 kJ/mol (Figure 4b). These results suggest that excessive binding strength forms a thermodynamic trap that hinders desorption. In terms of selectivity, the correlation profiles reveal a distinct asymmetry in Figure 4d–f. While high CO2 affinity defines the upper bound of selectivity (Figure 4d), the data in Figure 4e shows that C2H4 binding strength acts as a critical threshold. All highly selective MOFs are strictly confined to a low-enthalpy region (Q0st(C2H4) < 30 kJ/mol), beyond which selectivity suffers a sharp attenuation due to the competitive occupancy of adsorption sites by C2H4. While O2 affinity plays a negligible role (Figure 4f), the performance ceiling depends on a trade-off. Specifically, CO2 attraction must be maximized while a rigorous low-affinity boundary is maintained for C2H4 [40,41]. Synthesizing these factors, the composite TSN metric in Figure 4g,h identifies a clear optimal spot centered at Q0st(CO2) 42.7 kJ/mol and Q0st(C2H4) between 15 and 24 kJ/mol. This threshold-driven behavior indicates that enhanced CO2 affinity defines the performance ceiling. However, the minimization of C2H4 interference acts as the decisive limiting factor for practical viability, while O2 adsorption plays only a negligible secondary role (Figure 4i).
Figure 4.
Correlations between performance metrics and infinite-dilution isosteric heats of adsorption Q0st. (a–c) Working capacity (ΔN) versus Q0st for CO2, C2H4, and O2, respectively. (d–f) lg(S) versus Q0st for CO2, C2H4, and O2, respectively. (g–i) TSN versus Q0st for CO2, C2H4, and O2, respectively. The top 15 MOFs are highlighted as red points, and the remaining MOFs are shown in gray points.
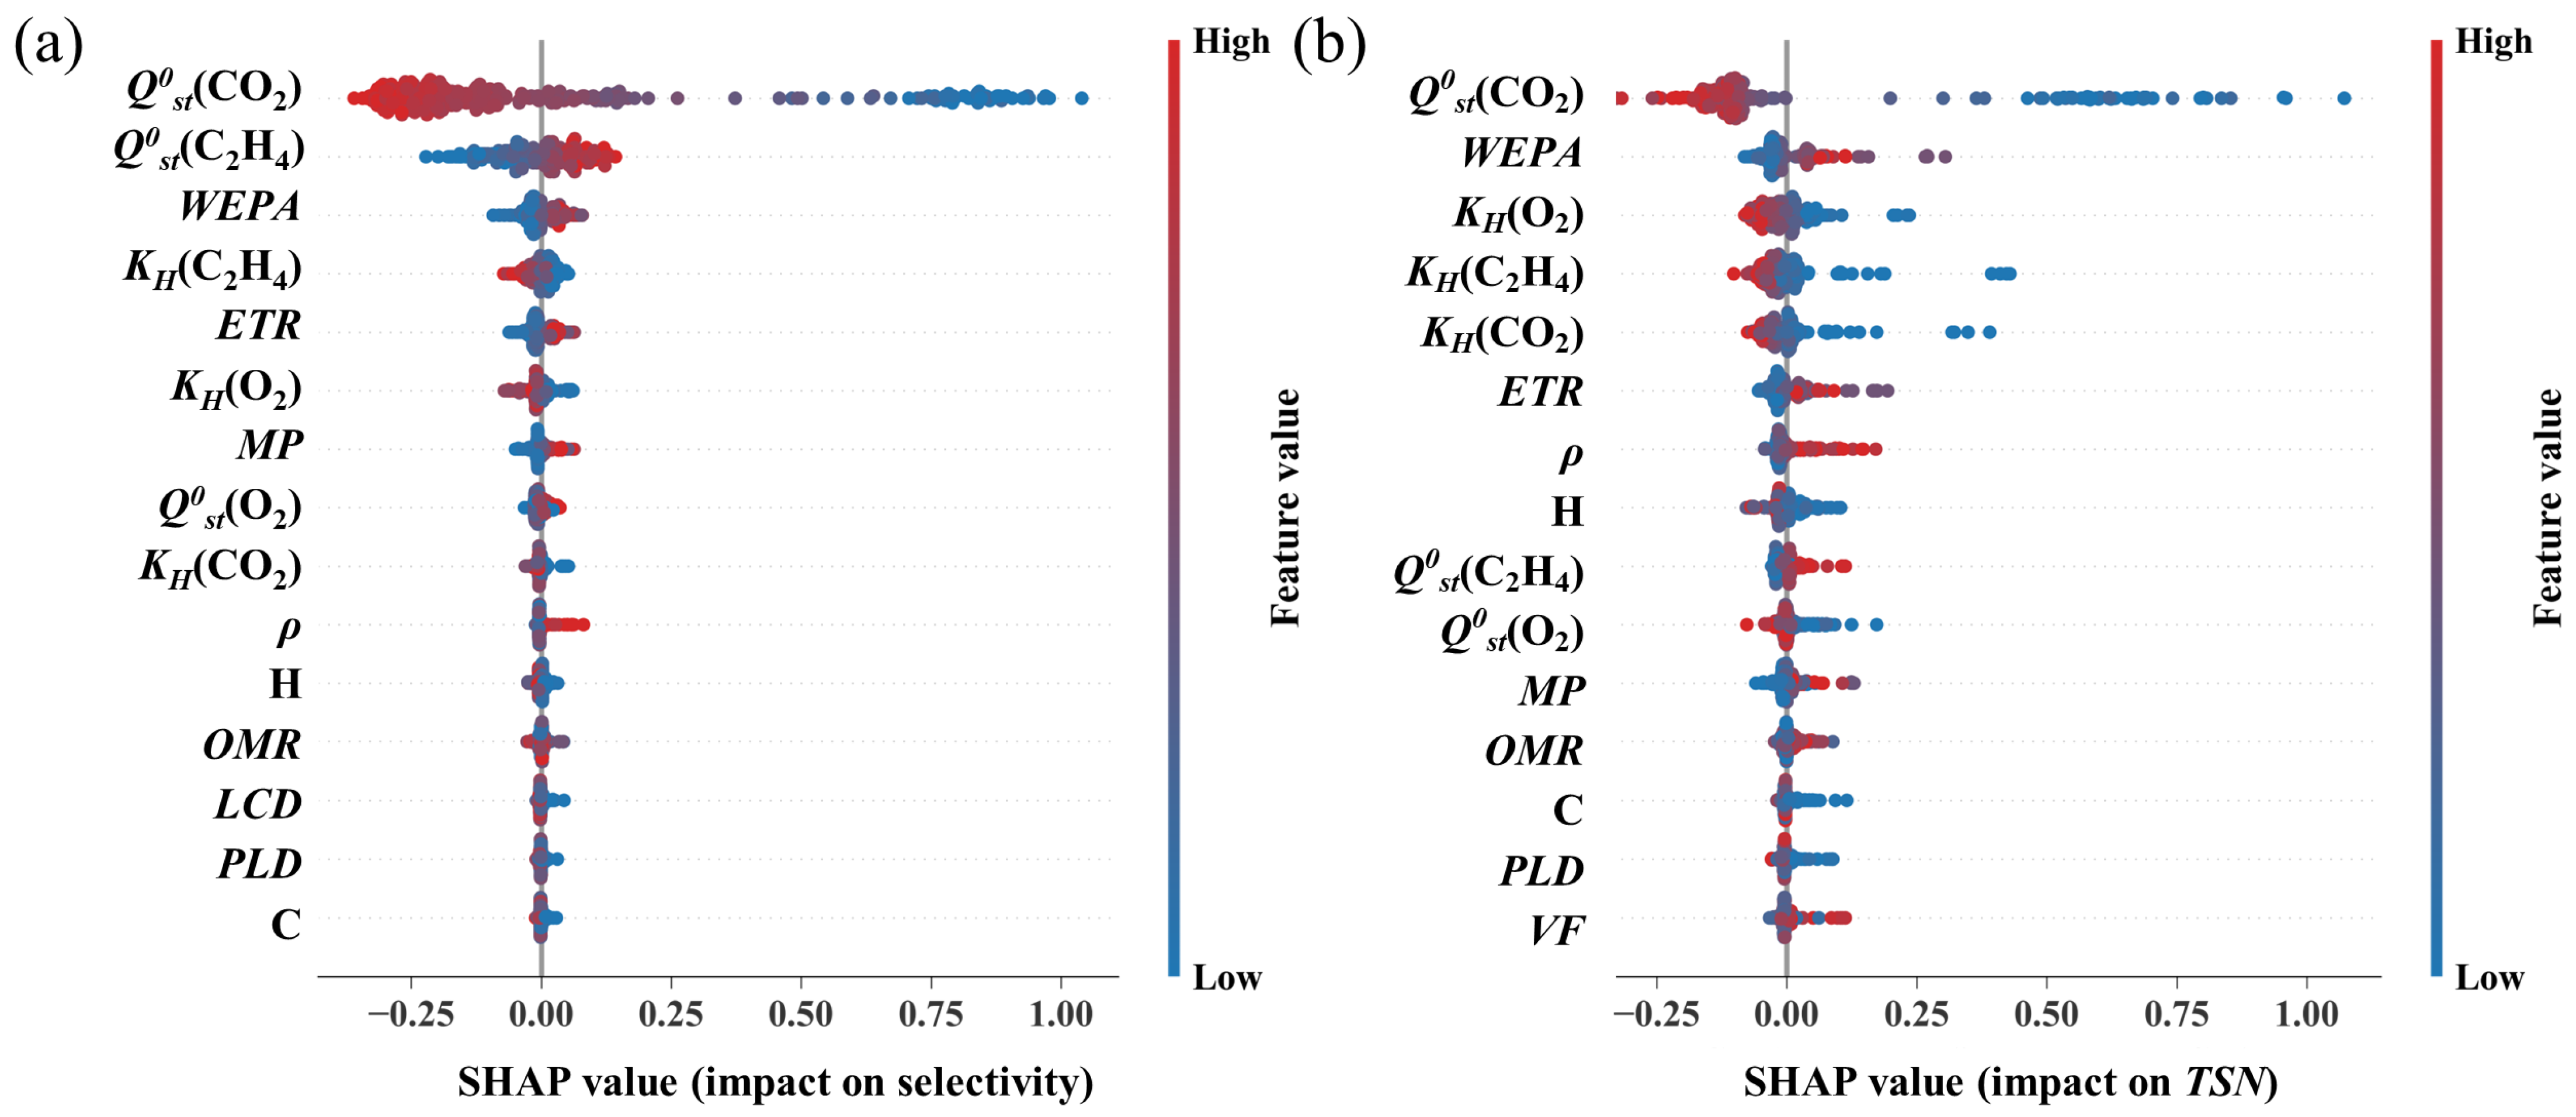
SHAP analysis further elucidates the relative contribution of each descriptor. As shown in Figure 5a, the feature ranking identifies Q0st(CO2), Q0st(C2H4), WEPA, KH(C2H4), and ETR as the dominant drivers governing selectivity. Specifically, Q0st descriptors play a decisive role in the competitive landscape. High Q0st(CO2) values correspond to strongly positive SHAP contributions, confirming that robust adsorbate-framework interactions enable CO2 to dominate adsorption sites. Conversely, elevated Q0st(C2H4) values shift the SHAP distribution toward the negative regime, indicating that heightened ethylene affinity severely undermines discriminatory power. This mechanism is reinforced by the KH descriptors: minimizing the baseline affinity for competitors KH(C2H4) and KH(O2) is critical for maintaining a CO2-selective environment, particularly at low partial pressures.
Figure 5.
SHAP value distributions for (a) selectivity and (b) TSN based on the RF model.
Electronic descriptors WEPA and ETR jointly govern the polarity-related characteristics of MOF frameworks at different structural scales and exert a dominant influence on CO2 selectivity [42,43]. WEPA reflects the overall electronegativity background of the framework, determined by the averaged electron-withdrawing contributions of its constituent atoms. Higher WEPA values reflect frameworks enriched with electronegative atoms (O, N, F). These increase global polarity and strengthen electrostatic interactions with the CO2 quadrupole. In contrast, ETR characterizes the density of electronegative atoms acting as localized polar sites. These create concentrated electric fields that promote directional electrostatic or Lewis acid-base interactions with CO2. Owing to the much weaker electrostatic response of C2H4, these highly polar environments preferentially amplify CO2 binding and thus enhance selectivity [44].
Regarding the composite metric TSN, Q0st(CO2) emerges as the primary determinant and exerts a profound positive influence (Figure 5b). Intensifying MOF-CO2 interactions simultaneously amplifies both adsorption capacity and selectivity, thereby compounding their effect on the final TSN value. Similarly, WEPA contributes positively by selectively enhancing the capture of quadrupolar CO2. In contrast, KH(O2) and KH(C2H4) display strong negative correlations, where minimized affinity for non-target species is essential to preserve TSN. A counterintuitive finding is the negative influence of KH(CO2), with optimal TSN scores clustering at moderate-to-low values. Extremely high KH(C2H4) can indicate ultra-strong sites that attract polarizable C2H4, creating a competitive binding effect that reduces overall selectivity.
3.5. Mechanistic Insights and Structural Feasibility of Top Candidates
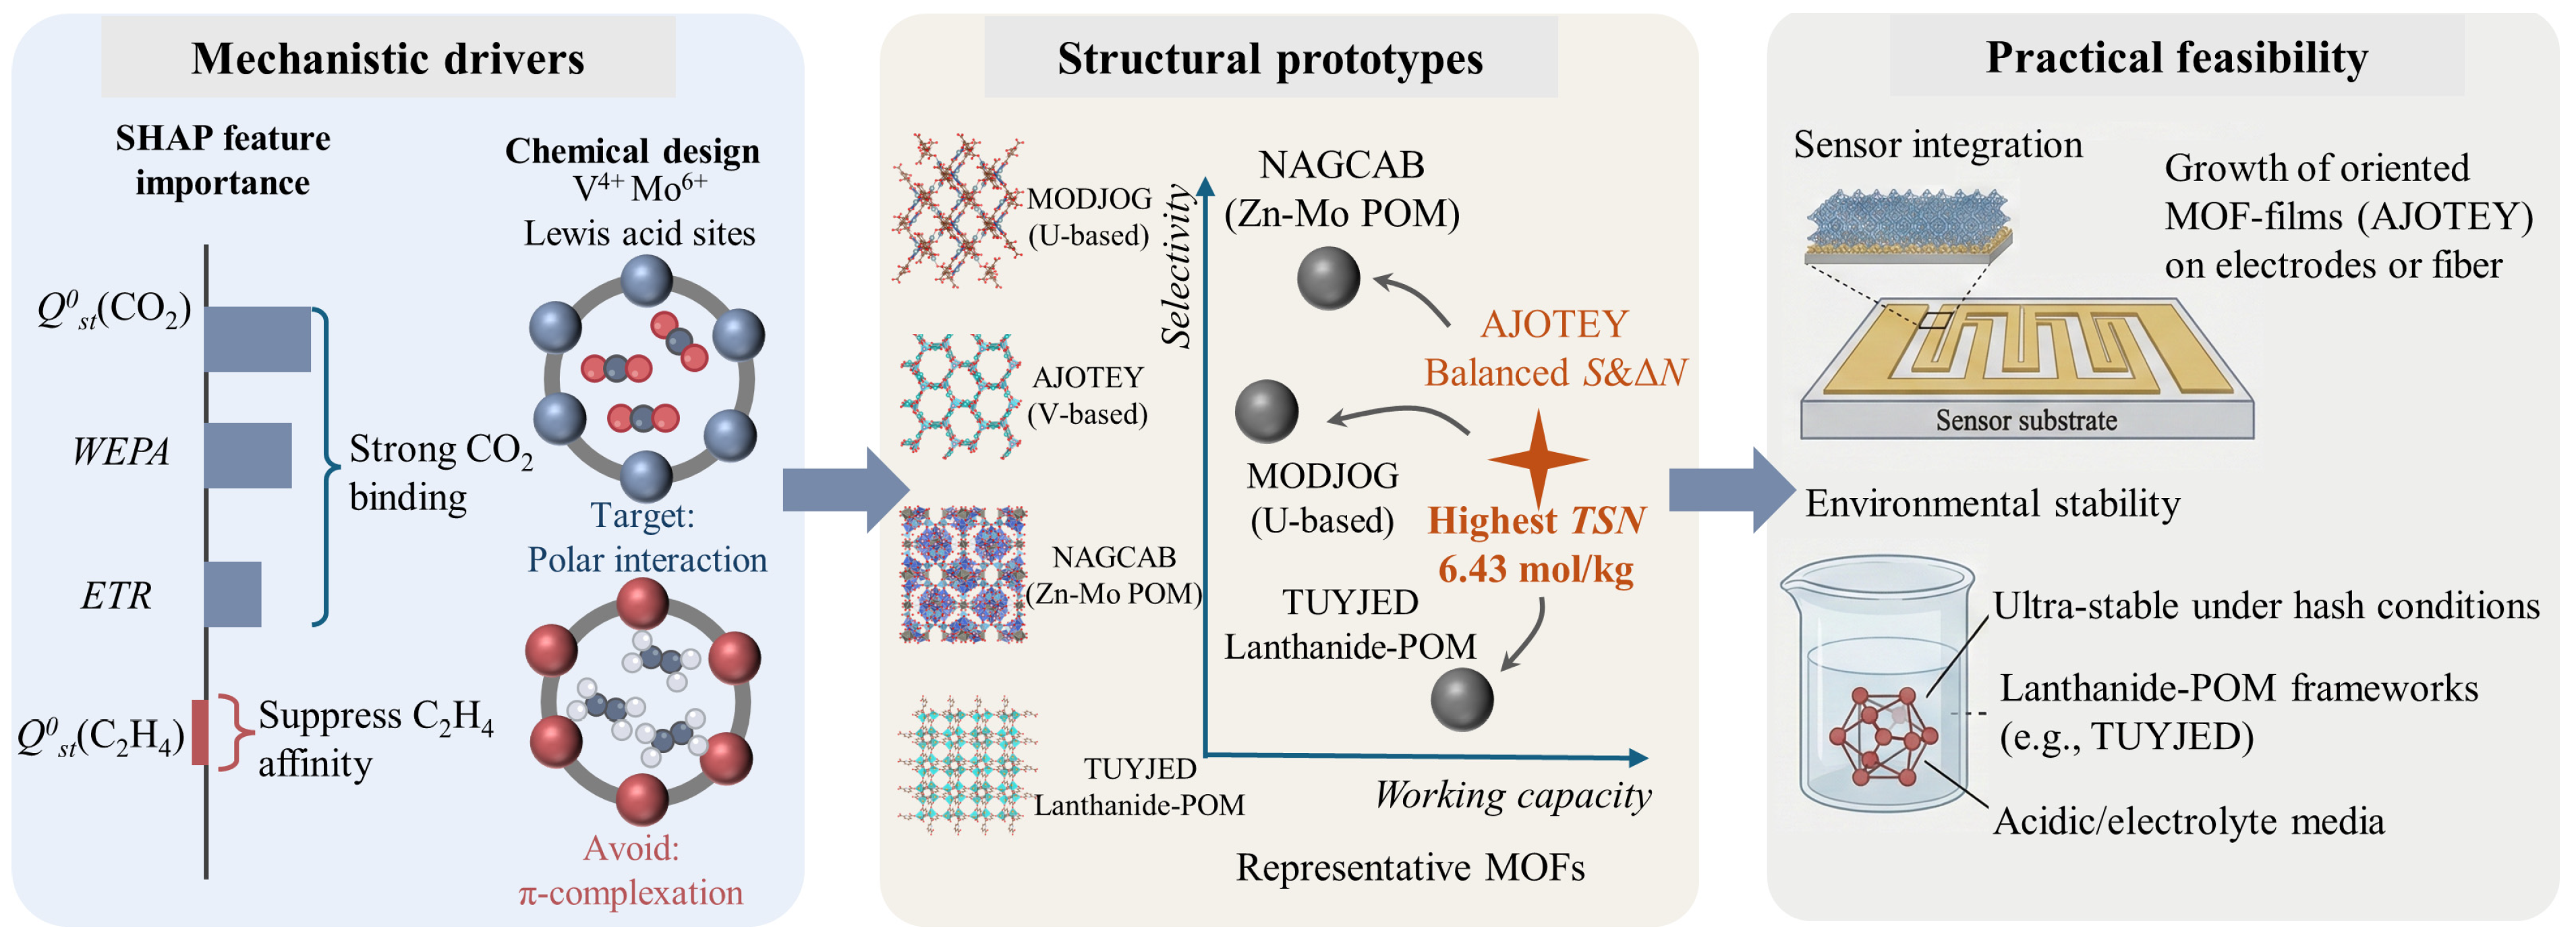
Consistent with physical interpretations, SHAP value analysis reveals that optimal MOFs must simultaneously strengthen CO2 adsorption while disfavoring C2H4 uptake. High positive SHAP contributions from Q0st(CO2), WEPA, and ETR confirm that a polar framework provides the fundamental basis for strong CO2 binding (Figure 6). In contrast, the strongly negative SHAP values of Q0st(C2H4) indicate that suppressing the affinity for competing C2H4 is equally critical. Chemically, this selective profile is achieved by a specific design strategy: the absence of strong π-complexation sites. The top candidates deliberately avoid low-valent transition metals (e.g., Cu+, Ag+) and high-density aromatic linkers that could enable π-π stacking with ethylene [40,45,46]. Instead, they predominantly utilize high-valent metal centers (e.g., V4+, U6+, Mo5+/6+, W6+) and lanthanides (e.g., Sm3+, Nd3+). As hard Lewis acid centers within inorganic or POM clusters, these metals optimize CO2 quadrupole interactions while suppressing affinity for non-polar O2 and weakly polar C2H4 [41,47].
Figure 6.
Mechanistic, structural, and practical determinants of high-performance MOF candidates for selective CO2 recognition.
This balance is perfectly exemplified by AJOTEY, the top-performing candidate (TSN = 6.43 mol/kg) [48]. Structurally, AJOTEY is a V4+-based framework constructed from vanadium-oxo clusters and phosphito linkers. The high density of polar V=O and V-OH sites serves as directional CO2 binding centers [49]. Geometrically, it aligns with the identified optimal profile, featuring a PLD of 5.20 Å, LCD of 5.91 Å, moderate ASA of 1245.70 m2/g, and high VF (0.60). Statistical validation across the top 15 candidates reinforces the importance of this intermediate pore window (average PLD ~4.5 Å), which successfully balances molecular sieving effects with the accessibility required for robust gas transport.
The trade-off between selectivity and capacity is evident when comparing specific candidates. NAGCAB (a Zn-Mo POM-based MOF) achieves the highest selectivity of 49.10 but ranks only second in TSN due to its modest working capacity (Tables S4 and S5). Its relatively small pores (PLD 6.99 Å) strongly enhance selectivity through confinement but limit total uptake. Conversely, AJOTEY balances both metrics by sacrificing some selectivity to achieve nearly double the working capacity (Figure 6). Similarly, NAQRAA and NAQREE (Sm/Eu-based POM-MOFs) adopt smaller PLD values (~4.2 Å) with larger LCD (~8.8 Å), creating elongated pore geometries that sustain high selectivity while preserving essential void space [50,51,52]. In contrast, MODJOG (a U-based framework) maximizes selectivity at the expense of capacity, resulting in a lower TSN. These comparisons demonstrate the model’s efficacy in prioritizing materials that resolve inherent adsorption trade-off.
Finally, regarding practical feasibility, all 15 top candidates are experimentally synthesizable, as evidenced by the CSD. Film formation is particularly promising for sensor integration (Figure 6). Many of these structures, including AJOTEY and POM-based families (like NAGCAB and NAQRAA), offer unique electrostatic properties suitable for deposition on sensor electrodes [49,53]. In terms of stability, the Lanthanide-POM based candidates such as TUYJED (Nd/W-based) and the LEYREO series (Pr/Mo/V-based) provide a robust alternative [52,53]. Rigid polyoxometalate backbones ensure structural integrity in acidic and oxidative environments, offering superior resistance to humidity and electrolyte leakage. Thus, the screened library offers a versatile toolkit: V-based AJOTEY as the high-performance standard, and Lanthanide/POM-MOFs as ultra-stable backups for harsh operating conditions.
4. Conclusions
In summary, this research demonstrates an efficient methodology for screening MOFs for early CO2 recognition during lithium-ion battery thermal runaway. By integrating GCMC simulations with an RF model, we successfully identified optimal materials capable of distinguishing CO2 from C2H4 and O2 under competitive conditions. SHAP analysis reveals that performance is governed by a delicate balance of adsorption energetics and pore accessibility. Specifically, CO2 selectivity originates from interaction strength differences driven by hard Lewis acid centers and polar inorganic clusters rather than simple size exclusion. Our findings indicate that high-performing MOFs do not possess extreme structural traits but instead fall within a narrow, balanced design window: moderate ASA (965–1975 m2/g), constrained pore openings (PLD ≈ 4–7 Å), high VF (>0.6), and low framework ρ (<1.3 g/cm3). Among the candidates, AJOTEY stands out by resolving the inherent capacity-selectivity trade-off, achieving a superior TSN value (6.43 mol/kg). Furthermore, the identification of Lanthanide-POM based frameworks provide versatile ultra-stable alternatives suitable for the harsh chemical environments of battery packs. This computational workflow significantly accelerates the discovery of sensing materials and establishes a reliable foundation for developing highly selective CO2 sensors to enhance battery safety systems.
Supplementary Materials
The following supporting information can be downloaded at: https://www.mdpi.com/article/10.3390/nano16040245/s1, Table S1: Descriptors characterizing structural and chemical properties of MOFs; Table S2: Calculation of other chemical descriptors; Table S3: The prediction scores of ML models for individual metrics; Table S4: Basic properties and separation performances of the fifteen top-performing MOFs sorted by selectivity; Table S5: Basic properties and separation performances of the fifteen top-performing MOFs sorted by TSN; Figure S1: Scatter plots showing the relationship between working capacity (ΔN) and five key structural descriptors (ASA, PLD, LCD, ρ, VF). The top 15 MOFs are highlighted as red points, and the remaining MOFs are shown in gray points.
Author Contributions
Conceptualization and Supervision and Writing—original draft and Formal analysis and Funding acquisition: X.W. (Xian Wei); Data curation and Investigation and Visualization: X.L. (Xin Li); Data curation and Formal analysis: X.W. (Xiong Wang); Resources and Writing—review and editing: X.L. (Xiaoyan Liu) and C.Z. (Chen Zhu). All authors have read and agreed to the published version of the manuscript.
Funding
This research was funded by the National Natural Science Foundation of China No. 62405141.
Data Availability Statement
The raw data supporting the conclusions of this article will be made available by the authors on request.
Acknowledgments
During the preparation of this work, the authors used ChatGPT 5.0 in order to improve language. After using this tool, the authors reviewed and edited the content as needed and take full responsibility for the content of the publication.
Conflicts of Interest
The authors declare no conflicts of interest.
References
- Grey, C.P.; Tarascon, J.M. Sustainability and in Situ Monitoring in Battery Development. Nat. Mater. 2016, 16, 45–56. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Wei, G.; Huang, R.; Zhang, G.; Jiang, B.; Zhu, J.; Guo, Y.; Han, G.; Wei, X.; Dai, H. A Comprehensive Insight into the Thermal Runaway Issues in the View of Lithium-Ion Battery Intrinsic Safety Performance and Venting Gas Explosion Hazards. Appl. Energy 2023, 349, 121651. [Google Scholar] [CrossRef] [Scilit]
- Gulsoy, B.; Chen, H.; Briggs, C.; Vincent, T.A.; Sansom, J.E.H.; Marco, J. Real-Time Simultaneous Monitoring of Internal Temperature and Gas Pressure in Cylindrical Cells During Thermal Runaway. J. Power Sources 2024, 617, 235147. [Google Scholar] [CrossRef] [Scilit]
- Zhang, J.; Li, F.; Yu, L.; Wang, Y.; Wang, K.; Chang, C.; Li, M.; Hao, W.; Qian, X. Research on the Explosive Characteristics and Suppression Mechanisms of Gas Generation During Thermal Runaway of Batteries in a Charged State. Chem. Eng. J. 2025, 505, 159699. [Google Scholar] [CrossRef] [Scilit]
- Lupan, O.; Ababii, N.; Mishra, A.K.; Gronenberg, O.; Vahl, A.; Schürmann, U.; Duppel, V.; Krüger, H.; Chow, L.; Kienle, L.; et al. Single CuO/Cu2O/Cu Microwire Covered by a Nanowire Network as a Gas Sensor for the Detection of Battery Hazards. ACS Appl. Mater. Interfaces 2020, 12, 42248–42263. [Google Scholar] [CrossRef] [Scilit]
- Cai, T.; Valecha, P.; Tran, V.; Engle, B.; Stefanopoulou, A.; Siegel, J. Detection of Li-Ion Battery Failure and Venting with Carbon Dioxide Sensors. eTransportation 2021, 7, 100100. [Google Scholar] [CrossRef] [Scilit]
- Wang, Z.; Zhu, L.; Zhang, J.; Wang, J.; Cui, X.; Chen, X.; Liu, W.; Ma, H.; Wang, J.; Yan, W. Zif-8 Loaded Ag/Zno Electrospun Nanofibers Enabling High-Performance H2 Gas Sensing for Battery Safety Early Warning. Sens. Actuators B Chem. 2024, 418, 136276. [Google Scholar] [CrossRef] [Scilit]
- Ren, Q.; Zhang, X.; Guo, Y.; Xu, M.; Zhu, H.; Yun, J.; Zhao, W.; Zhang, Z.; Wang, Y. Shape-Controlled Sno and Their Improved Properties in the Field of Gas Sensor, Photocatalysis, and Lithium-Ion Battery. Sens. Actuators B Chem. 2022, 372, 132622. [Google Scholar] [CrossRef] [Scilit]
- Wang, X.-X.; Li, Q.-T.; Zhou, X.-Y.; Hu, Y.-M.; Guo, X. Monitoring Thermal Runaway of Lithium-Ion Batteries by Means of Gas Sensors. Sens. Actuators B Chem. 2024, 411, 135703. [Google Scholar] [CrossRef] [Scilit]
- Sheng, S.; Li, H.; Zhang, Y.; Li, L.; Xiao, K.; Huang, X.; Liu, Y.; Xu, W.; Li, Z.; Yan, L.; et al. Early Detection of Lithium Battery Lakage Using a Highly Sensitive in Situ Zif-8 Membrane-Coated Micro-Nano Optical Fibre. Light Adv. Manuf. 2025, 6, 130–141. [Google Scholar]
- Wang, X.; Zhang, T. An Overview: A Design Strategy for Dioxide Carbon Qcm Gas Sensor Based on Mofs or Cofs. Adv. Sensor Res. 2025, 4, 2400170. [Google Scholar] [CrossRef] [Scilit]
- Yanez-Aulestia, A.; Trejos, V.M.; Esparza-Schulz, J.M.; Ibarra, I.A.; Sanchez-Gonzalez, E. Chemically Modified Hkust-1(Cu) for Gas Adsorption and Separation: Mixed-Metal and Hierarchical Porosity. ACS Appl. Mater. Interfaces 2024, 16, 65581–65591. [Google Scholar] [CrossRef] [Scilit]
- Zelenka, T.; Simanova, K.; Saini, R.; Zelenkova, G.; Nehra, S.P.; Sharma, A.; Almasi, M. Carbon Dioxide and Hydrogen Adsorption Study on Surface-Modified Hkust-1 with Diamine/Triamine. Sci. Rep. 2022, 12, 17366. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Mahajan, S.; Elfving, J.; Lahtinen, M. Evaluating the Viability of Ethylenediamine-Functionalized Mg-Mof-74 in Direct Air Capture: The Challenges of Stability and Slow Adsorption Rate. J. Environ. Chem. Eng. 2024, 12, 112193. [Google Scholar] [CrossRef] [Scilit]
- Qazvini, O.T.; Babarao, R.; Telfer, S.G. Selective Capture of Carbon Dioxide from Hydrocarbons Using a Metal-Organic Framework. Nat. Commun. 2021, 12, 197. [Google Scholar] [CrossRef] [Scilit]
- Qi, S.; Cheng, Y.; Li, Z.; Wang, J.; Li, H.; Zhang, C. Advanced Deep Learning Techniques for Battery Thermal Management in New Energy Vehicles. Energies 2024, 17, 4132. [Google Scholar] [CrossRef] [Scilit]
- Zuo, W.; Zheng, H.; He, T.; Vishwanath, V.; Chan, M.K.Y.; Stevens, R.L.; Amine, K.; Xu, G.-L. Large Language Models for Batteries. Joule 2025, 9, 102037. [Google Scholar] [CrossRef] [Scilit]
- Sung, I.T.; Cheng, Y.-H.; Hsieh, C.-M.; Lin, L.-C. Machine Learning for Gas Adsorption in Metal–Organic Frameworks: A Review on Predictive Descriptors. Ind. Eng. Chem. Res. 2025, 64, 1859–1875. [Google Scholar] [CrossRef] [Scilit]
- Gandhi, A.; Hasan, M.M.F. Machine Learning for the Design and Discovery of Zeolites and Porous Crystalline Materials. Curr. Opin. Chem. Eng. 2022, 35, 100739. [Google Scholar] [CrossRef] [Scilit]
- Daglar, H.; Keskin, S. Combining Machine Learning and Molecular Simulations to Unlock Gas Separation Potentials of Mof Membranes and Mof/Polymer Mmms. ACS Appl. Mater. Interfaces 2022, 14, 32134–32148. [Google Scholar] [CrossRef] [Scilit]
- Pilz, L.; Koenig, M.; Schwotzer, M.; Gliemann, H.; Wöll, C.; Tsotsalas, M. Enhancing the Quality of Mof Thin Films for Device Integration through Machine Learning: A Case Study on Hkust-1 Surmof Optimization. Adv. Funct. Mater. 2024, 34, 2404631. [Google Scholar] [CrossRef] [Scilit]
- Luo, Y.; Bag, S.; Zaremba, O.; Andreo, J.; Wuttke, S.; Tsotsalas, M.; Friederich, P. Mof Synthesis Prediction Enabled by Automatic Data Mining and Machine Learning. Angew. Chem. Int. Ed. 2021, 61, e202200242. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Qiao, Z.; Xu, Q.; Jiang, J. Computational Screening of Hydrophobic Metal–Organic Frameworks for the Separation of H2s and CO2 from Natural Gas. J. Mater. Chem. A 2018, 6, 18898–18905. [Google Scholar] [CrossRef] [Scilit]
- Wang, Y.; Jiang, Z.J.; Lu, W.; Li, D. Machine Learning-Assisted Exploration of Chemical Space of Mof-5 Analogs for Enhanced C2H6/C2H4 Separation. Angew. Chem. Int. Ed. 2025, 64, e202500783. [Google Scholar] [CrossRef] [Scilit]
- Orhan, I.B.; Daglar, H.; Keskin, S.; Le, T.C.; Babarao, R. Prediction of O2/N2 Selectivity in Metal–Organic Frameworks Via High-Throughput Computational Screening and Machine Learning. ACS Appl. Mater. Interfaces 2021, 14, 736–749. [Google Scholar] [CrossRef] [Scilit]
- Yang, Y.; Shin, Y.K.; Ooe, H.; Hasegawa, U.; Yamane, S.; Yamada, H.; van Duin, A.C.T.; Murase, Y.; Mauro, J.C. Adsorption of CO2 by Amine-Functionalized Metal–Organic Frameworks Using Gcmc and Reaxff-Based Metadynamics Simulations. J. Phys. Chem. C 2024, 128, 5257–5270. [Google Scholar] [CrossRef] [Scilit]
- Parsaei, M.; Akhbari, K.; Tylianakis, E.; Froudakis, G.E.; White, J.M. Efficient Gas Adsorption in Mut-11: Insights from Theoretical Calculations and Gcmc Simulations. Cryst. Growth Des. 2024, 24, 10299–10313. [Google Scholar] [CrossRef] [Scilit]
- Sadeghi, M.; Esmaeilzadeh, F.; Mowla, D.; Zandifar, A. Improving CO2 Capture in Utsa-16(Zn) Via Alkali and Alkaline Earth Metal Introduction: Gcmc and Md Simulations Study. Sep. Purif. Technol. 2024, 338, 126534. [Google Scholar] [CrossRef] [Scilit]
- Wang, Y.; Jiang, Z.-J.; Wang, D.-R.; Lu, W.; Li, D. Machine Learning-Assisted Discovery of Propane-Selective Metal–Organic Frameworks. J. Am. Chem. Soc. 2024, 146, 6955–6961. [Google Scholar] [CrossRef] [Scilit]
- Lim, Y.; Kim, B.; Kim, J. Data-Driven Design of Flexible Metal–Organic Frameworks for Gas Storage. Chem. Mater. 2024, 36, 5465–5473. [Google Scholar] [CrossRef] [Scilit]
- Pardakhti, M.; Moharreri, E.; Wanik, D.; Suib, S.L.; Srivastava, R. Machine Learning Using Combined Structural and Chemical Descriptors for Prediction of Methane Adsorption Performance of Metal Organic Frameworks (Mofs). ACS Comb. Sci. 2017, 19, 640–645. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Gao, W.; Zheng, W.; Yan, K.; Sun, W.; Zhao, L. Accelerating the Discovery of Acid Gas-Selective Mofs for Natural Gas Purification: A Combination of Machine Learning and Molecular Fingerprint. Fuel 2023, 350, 128757. [Google Scholar] [CrossRef] [Scilit]
- Wang, Z.; Zhu, L.; Liu, J.; Wang, J.; Yan, W. Gas Sensing Technology for the Detection and Early Warning of Battery Thermal Runaway: A Review. Energy Fuels 2022, 36, 6038–6057. [Google Scholar] [CrossRef] [Scilit]
- Rappé, A.K.; Casewit, C.J.; Colwell, K.S.; III, W.A.G.; Skiff, W.M. Uff, a Full Periodic Table Force Field for Molecular Mechanics and Molecular Dynamics Simulations. J. Am. Chem. Soc. 1992, 114, 10024–10035. [Google Scholar] [CrossRef] [Scilit]
- Dubbeldam, D.; Calero, S.; Ellis, D.E.; Snurr, R.Q. Raspa: Molecular Simulation Software for Adsorption and Diffusion in Flexible Nanoporous Materials. Mol. Simul. 2015, 42, 81–101. [Google Scholar] [CrossRef] [Scilit]
- Widom, B. Some Topics in the Theory of Fluids. J. Chem. Phys. 1963, 39, 2808–2812. [Google Scholar] [CrossRef] [Scilit]
- Wilmer, C.E.; Farha, O.K.; Bae, Y.-S.; Hupp, J.T.; Snurr, R.Q. Structure–Property Relationships of Porous Materials for Carbon Dioxide Separation and Capture. Energy Environ. Sci. 2012, 5, 9849. [Google Scholar] [CrossRef] [Scilit]
- Firooz, S.K.; Armstrong, D.W. Metal-Organic Frameworks in Separations: A Review. Anal. Chim. Acta 2022, 1234, 340208. [Google Scholar] [CrossRef] [Scilit]
- Turaeva, N.; Yablonsky, G.; Fushimi, R. Fermi-Based Kinetic Model for the Sabatier Reaction: Sabatier Principle and Beyond It. J. Phys. Chem. C 2025, 129, 7730–7739. [Google Scholar] [CrossRef] [Scilit]
- Yan, X.-W.; Bigdeli, F.; Abbasi-Azad, M.; Wang, S.-J.; Morsali, A. Selective Separation of CO2/CH4 Gases by Metal-Organic Framework-Based Composites. Coord. Chem. Rev. 2024, 520, 216126. [Google Scholar] [CrossRef] [Scilit]
- Sumida, K.; Rogow, D.L.; Mason, J.A.; McDonald, T.M.; Bloch, E.D.; Herm, Z.R.; Bae, T.H.; Long, J.R. Carbon Dioxide Capture in Metal-Organic Frameworks. Chem. Rev. 2012, 112, 724–781. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Wilmer, C.E.; Leaf, M.; Lee, C.Y.; Farha, O.K.; Hauser, B.G.; Hupp, J.T.; Snurr, R.Q. Large-Scale Screening of Hypothetical Metal-Organic Frameworks. Nat. Chem. 2011, 4, 83–89. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Smit, B.; Maesen, T.L. Towards a Molecular Understanding of Shape Selectivity. Nature 2008, 451, 671–678. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Laha, S.; Dwarkanath, N.; Sharma, A.; Rambabu, D.; Balasubramanian, S.; Maji, T.K. Tailoring a Robust Al-Mof for Trapping C2H6 and C2H2 Towards Efficient C2H4 Purification from Quaternary Mixtures. Chem. Sci. 2022, 13, 7172–7180. [Google Scholar] [CrossRef] [Scilit]
- Li, J.-R.; Ma, Y.; McCarthy, M.C.; Sculley, J.; Yu, J.; Jeong, H.-K.; Balbuena, P.B.; Zhou, H.-C. Carbon Dioxide Capture-Related Gas Adsorption and Separation in Metal-Organic Frameworks. Coord. Chem. Rev. 2011, 255, 1791–1823. [Google Scholar] [CrossRef] [Scilit]
- Li, B.; Zhang, Y.; Krishna, R.; Yao, K.; Han, Y.; Wu, Z.; Ma, D.; Shi, Z.; Pham, T.; Space, B.; et al. Introduction of Pi-Complexation into Porous Aromatic Framework for Highly Selective Adsorption of Ethylene over Ethane. J. Am. Chem. Soc. 2014, 136, 8654–8660. [Google Scholar] [CrossRef] [Scilit]
- Wang, C.; Yan, J.; Chen, S.; Liu, Y. High-Valence Metal-Organic Framework Materials Constructed from Metal-Oxo Clusters: Opportunities and Challenges. ChemPlusChem 2023, 88, e202200462. [Google Scholar] [CrossRef] [Scilit]
- Harrison, W.T.A. (CN3H6)2·(VO)3(H2O)3(HPO3)4·3H2O, a Guanidinium Vanadium Phosphite Hydrate Based on a (3,4) Connected Net: An “Orthogonal Templating” Effect? Solid State Sci. 2003, 5, 297–302. [Google Scholar] [CrossRef] [Scilit]
- Liu, Y.-Y.; Couck, S.; Vandichel, M.; Grzywa, M.; Leus, K.; Biswas, S.; Volkmer, D.; Gascon, J.; Kapteijn, F.; Denayer, J.F.M.; et al. New V(Iv)-Based Metal-Organic Framework Having Framework Flexibility and High CO2 Adsorption Capacity. Inorg. Chem. 2013, 52, 113–120. [Google Scholar] [CrossRef] [Scilit]
- Hou, Y.N.; Xing, Y.H.; Bai, F.Y.; Guan, Q.L.; Wang, X.; Zhang, R.; Shi, Z. Synthesis, Crystal Structure, Photoluminescence Property and Photoelectronic Behavior of Two Uranyl-Organic Frameworks Constructed from 1, 2, 4, 5-Benzenetetracarboxylic Acid as Ligand. Spectrochim. Acta A Mol. Biomol. Spectrosc. 2014, 123, 267–272. [Google Scholar] [CrossRef] [Scilit]
- Gao, Y.; Xu, Y.; Han, Z.; Li, C.; Cui, F.; Chi, Y.; Hu, C. Syntheses, Structures and Properties of 3d Inorganic–Organic Hybrid Frameworks Constructed from Lanthanide Polymer and Keggin-Type Tungstosilicate. J. Solid State Chem. 2010, 183, 1000–1006. [Google Scholar] [CrossRef] [Scilit]
- Perez-Botella, E.; Valencia, S.; Rey, F. Zeolites in Adsorption Processes: State of the Art and Future Prospects. Chem. Rev. 2022, 122, 17647–17695. [Google Scholar] [CrossRef] [Scilit]
- Wang, X.; Yue, L.; Zhou, P.; Fan, L.; He, Y. Lanthanide-Organic Frameworks Featuring Three-Dimensional Inorganic Connectivity for Multipurpose Hydrocarbon Separation. Inorg. Chem. 2021, 60, 17249–17257. [Google Scholar] [CrossRef] [Scilit]

Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2026 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license.