


Abstract
This study examines the influence of SO2 and its hydrate H2SO3 on the free energies of the core autocatalytic cycle of the formose reaction. We find that SO2 and H2SO3 readily condense with aldehyde and alcohol functional groups to form bisulfite analogs of formose proto-metabolites under modeled conditions. The bisulfite functional group can provide intramolecular catalytic enhancement in specific isomers towards aldol additions and the retroaldol step that regenerates two equivalents of glycolaldehyde from tetrose. The bisulfite moiety reduces the favorability of the parasitic Cannizzaro side-reaction both thermodynamically and kinetically, thus potentially furnishing more throughput towards forming sugars. As a prebiotic analog to phosphate, we find that bisulfite slightly stabilizes ribose over its C5 aldose diastereomers thermodynamically, although the effect is modest and may be influenced by solution dynamics.
1. Introduction
The formose reaction is frequently invoked when discussing the chemical origins of life. It is a potentially autocatalytic source of a wide range of sugars including ribose and glucose, key building blocks in extant biochemistry. Autocatalytic chemical cycles are ubiquitous in living organisms [1], where they are embedded in compartments and under tight regulation, thus resilient to fluctuating environmental conditions. Such cycles are rare in “simple” chemical reactions which can be heavily influenced by an ever-changing environment. This is one reason why the core formose cycle has garnered interest as a potential model system to examine how modern metabolic cycles emerged from the seemingly chaotic chemistry of a proto-metabolism.
The formose reaction begins with formaldehyde (CH2O). As a C1 species, it is the “food” molecule or building block for constructing larger molecules, but requires an initiator to jumpstart oligomerization. The reaction is messy: CH2O oligomerizes through a series of aldol reactions generating sugars (CH2O)n of increasing size and diversity [2], but generates side-products such as carboxylic acids and polyols via parasitic Cannizzaro reactions. While the reaction mechanisms have been examined experimentally [3,4,5], there are many intricacies where varying experimental conditions result in different product distributions, which is why this complex system of reactions continues to be examined [6,7,8,9,10,11,12]. Several computational groups that have written code to model large-scale systems chemistry [13,14,15,16] chose to examine the formose reaction as a test case, highlighting its intriguing complexity.
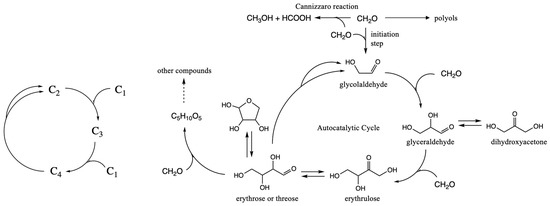
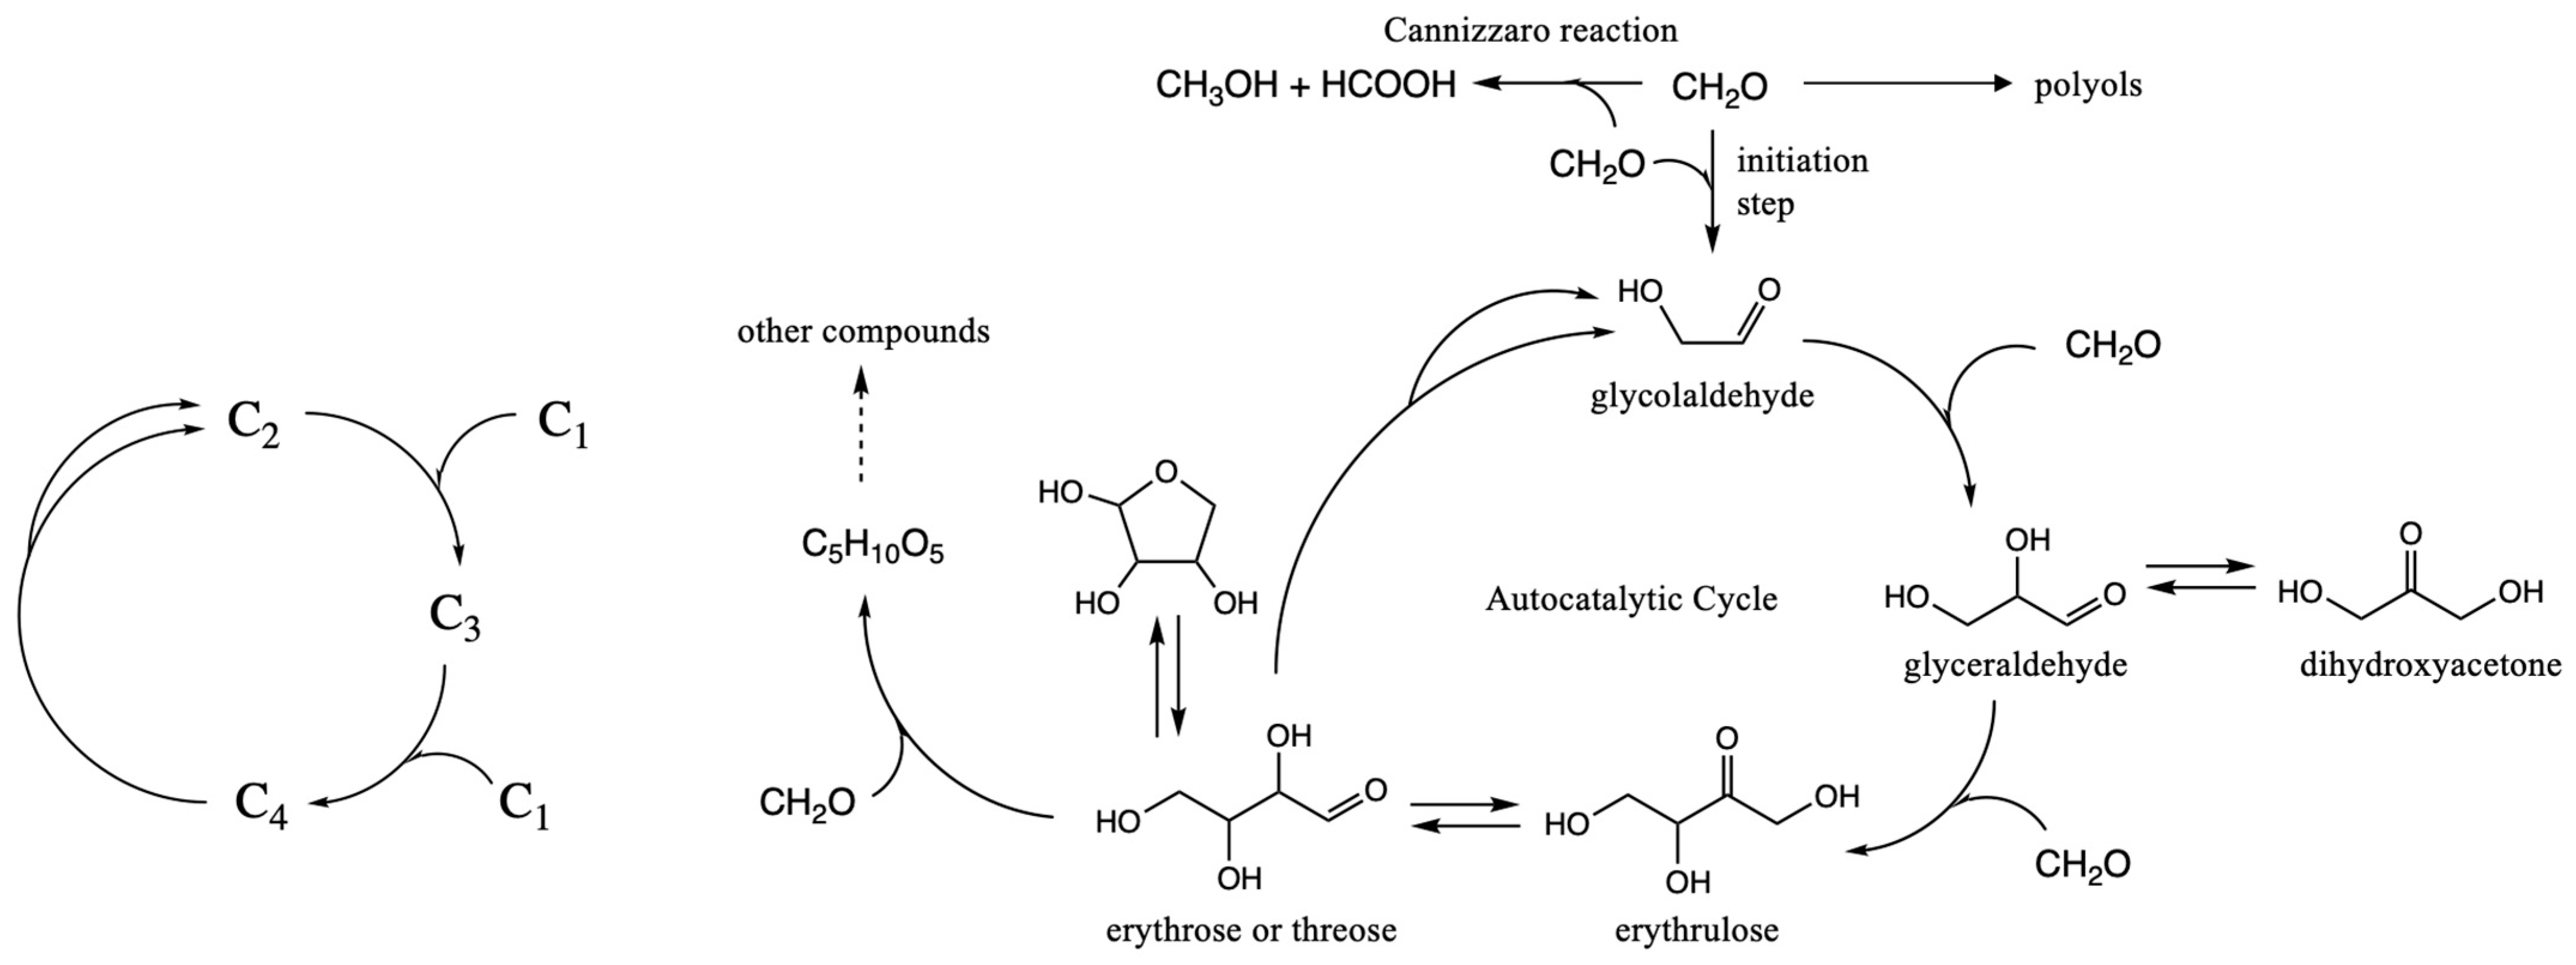
The core of the autocatalytic cycle nominally involves just three steps, represented by the simple abstract cycle on the left in Figure 1. The actual species involved are (CH2O)n molecules (n = 1–4), shown more explicitly on the right in Figure 1. If only CH2O is present initially, forming glycolaldehyde (C2H4O2) is very slow because C–C bond formation between two electrophilic carbons has a very high barrier [17]. However, if a small amount of this linchpin C2 species is formed or added as an initiator, the reaction proceeds rapidly via aldol reactions since an enol is now available as a nucleophile. The C2 glycolaldehyde reacts with CH2O to form C3 glyceraldehyde, which reacts further with CH2O to form C4 erythrulose. Autocatalysis is triggered if the C4 ketose isomerizes to an aldose which then generates two C2 molecules in a retro-aldol reaction; the increasing concentration of linchpin C2 rapidly eats up the C1 food as the cycle accelerates. The net reaction of this cycle is C1 + C1 → C2, not through the difficult direct dimerization, but rather facilitated by the C2, C3 and C4 species in a cycle of reactions. Larger sugars are formed through subsequent aldol reactions. Figure 1 shows one example (C4 + C1 → C5) but there are many other possibilities; we will explore the C2 + C3 → C5 reaction later in this article. While Figure 1 shows one example of the Cannizzaro reaction, note that all sugars present can potentially disproportionate into acids and polyols.
Figure 1.
Core autocatalytic cycle of the formose reaction. Reproduced from our previous work [18] under the terms and conditions of the Creative Commons Attribution (CC BY) 4.0 license.
Extant biochemistry utilizes only a small subset of sugars while the formose reaction generates hundreds in an unregulated synthesis. Ribose, a crucial building block required in the RNA World hypothesis [19], is produced in very low yield under standard alkaline conditions of the formose reaction; it rarely accumulates even in the presence of catalysts. Many early experimental studies focused on tweaking the environmental conditions and adding catalysts/co-substrates to boost the yield of ribose; examples include borate [20], cyanamide [21], silicates [22], hydroxyapatite [23], and various metal catalysts [24]. Of particular interest to our present work is phosphate: Eschenmoser’s group found that the aldol addition of the glycolaldehyde phosphate and glyceraldehyde phosphate yielded ribose (with phosphates attached) in high yield over its diastereomers [25]. Subsequent work in the Lane group using acetylphosphate under formose conditions also favored ribose stabilization [26]. Phosphates play a critical role in extant life as evidenced by the ubiquity of kinases, enzymes that control biochemical reactions by phosphorylating and dephosphorylating a wide range of substrates.
However, one challenge is that under the reducing conditions of Hadean earth, soluble phosphate was rare [27,28]; although a recent suggestion is that dissolved Fe2+ may increase the availability of soluble phosphate [29]. On the other hand, the atmosphere of the early earth may have been rich with sulfur. A study of Hadean zircons suggests that redox conditions corresponded to the fayalite-magnetite-quartz buffer [30], thus implying that volcanic activity generated SO2, which hydrates to form H2SO3 when dissolved in aqueous solution. As a weak acid, H2SO3 generates bisulfite (and sulfite). Benner and co-workers prop [31] that prebiotic chemistry involving bisulfite adducts likely could have occurred given the production of SO2 from Hadean volcanoes. They also provide experimental evidence that formaldehyde-bisulfite adducts are stable, effectively sequester CH2O and reduce the parasitic Cannizzaro side reactions [32] in the formose system. Thus, we hypothesize that the presence of bisulfite adducts may provide some form of selectivity towards ribose and other biologically-relevant sugars.
This article explores the role of bisulfite by calculating the thermodynamics and kinetics of the early steps in the formose reaction in the presence of bisulfite adducts. We previously used a similar approach to study if thiols provide selectivity in the formose reaction [18]: Briefly, our results suggested that a key barrier for the C2 + C1 → C3 addition reaction was modestly lowered, which led to a pathway resulting in a marginally exergonic C4 → C2 + C2 retro-aldol reaction (but with ΔG very close to zero). In the absence of the thiol group, this retro-aldol reaction is slightly endergonic, which limits its autocatalytic potential. However the messiness and complexity of formose steps remained as isomeric reactions with the thiol in different positions led to a wide range of products, which does not tame the overall selectivity problem.
Could bisulfite analogs provide more favorable energetics to initiate autocatalysis? Our present work quantifies free energy changes (and associated activation barriers) for bisulfite adduct formation, aldol reactions of adducts in the formose cycle, and competing Cannizzaro side-reactions. We also investigate if sulfite could play the analogous role to phosphate in selecting ribose in the C2 + C3 → C5 aldol reaction. Computational studies such as ours are a static model system for a set of baseline conditions, and therefore our calculated free energies will not necessarily correspond to the dynamic conditions of an actual formose system under experimental conditions. Before getting into the results and discussion, we describe our computational protocol and its limitations.
2. Computational Methods
We use a similar computational protocol to our recent work on the formose cycle [33] and its thiol analogs [18]. This allows us to make direct comparisons and extend our free energy map. Here, we provide a brief description of that protocol for the convenience of our readers. Much of the text in this section (and the first two paragraphs of the next section) is reproduced from those two articles (also published in this journal) since we think the description is both clear and succinct. Our calculated free energies using this quantum chemistry protocol showed good agreement with available experimental results for CHO systems [17,34,35,36].
The computational details are as follows: The geometry of each molecule is optimized and its electronic energy calculated at the B3LYP (Becke3-Lee-Yang-Parr) [37,38,39,40] flavor of density functional theory with the 6-311G** basis set. To maximize the probability of finding global minima, multiple conformers are generated. The optimized structures are embedded in a continuum dielectric to calculate the aqueous solvation contribution to the free energy. While this does not provide a specific concentration, it assumes a dilute solution such that the electrostatic field generated by a neighboring solute molecule is effectively screened by a dielectric representing water using the SMD (Solvation Model based on Density) implicit solvent [41].
Zero-point energy corrections are included, and we apply the standard temperature-dependent enthalpy correction term (for 298.15 K) from statistical mechanics by assuming translational and rotational corrections are a constant times kT, and that low frequency vibrational modes generally cancel out when calculating enthalpy differences. However, entropic corrections in aqueous solution are problematic [42,43,44]. Changes in free energy terms for translation and rotation are poorly defined in solution due to restricted complex motion, particularly as the size of the molecule increases (thus increasing its conformational entropy). Free energy corrections come from two different sources: thermal corrections and implicit solvent. Neither of these parameters is easily separable, nor do they constitute all the required parts of the free energy. We follow the approach of Deubel & Lau [45], assigning the solvation entropy of each species as half its gas-phase entropy (calculated using standard statistical mechanics approximations similar to the enthalpy calculations described above), based on proposals by Wertz [46] and Abraham [47] that upon dissolving in water, molecules lose a constant fraction (~0.5) of their entropy.
To estimate activation energies, transition states were optimized by including several explicit water and/or catalytic molecules to facilitate proton transfer. All calculated transition states have one significant negative eigenvalue corresponding to the reaction coordinate. Each transition state was checked to ensure it had the correct eigenvectors for bond breaking/forming connecting the transition state to its reactants and products. Several conformers are tested, and for each reaction we only report the lowest calculated barriers with the optimum number of solvent molecules added to assist in proton transfer.
It is necessary to validate one’s computational protocol with available experimental values. Our protocol performs well comparing the equilibrium concentrations in a self-oligomerizing solution of 1 M glycolaldehyde at 298 K to subsequent NMR measurements [35]. Our relative Gibbs free energies in aqueous solution are typically within 0.5 kcal/mol/mol compared to experiment. That being said, our protocol has systematic errors of 2–3 kcal/mol/mol when calculating barriers involving carbonyl chemistry when compared to experimental results. Going to a higher level of theory does not reduce this error [48]; nor does using anionic species; our protocol shows better agreement with experimental results for the neutral forms [36]. Quantum chemistry is about error cancellation, and our protocol (with neutral molecules, implicit solvent, and a simplistic entropy correction) has worked well in the CHO systems we have studied even with a systematic error for activation barriers.
3. Results and Discussion
Throughout this paper, the thermodynamics of a chemical reaction are denoted by ΔG. For transition states, kinetic barriers are denoted by ΔG‡ and will refer to the forward barrier unless otherwise specified. A consistent set of reference compounds allows us to globally compare energies. In several Figures, Grel values are found under each compound for local minima and in square brackets under an arrow for transition states. For a chemical reaction, ΔG = Grel(products) − Grel(reactants), and ΔG‡ compares Grel of the transition state to either the reactants or products depending on whether the forward or reverse reaction is being discussed. The Grel values for all compounds parsed into their separate enthalpic, entropic, and solvation free energy are provided in Supplementary Materials.
To connect this work to our prior thermodynamic map examining the formose cycle with H2S [18], we use a similar set of reference compounds: CO2, H2, H2O and H2SO3 are assigned a relative free energy, Grel of 0.0 kcal/mol. Instead of H2S as our reference for thiols, we use the hydrate of SO2 as our reference for bisulfite analogs. Using the hydrate provided a cleaner comparison, and also better matched experimental thermodynamic values in previous work [36]. The Grel values of all other species can be determined by calculating the change in free energy for forming the species, analogous to a free energy of formation. For example, the formation reaction of the bisulfite adduct of glycolaldehyde (C2H4SO4) is written as
2 CO2 + 4 H2 + H2SO3 → C2H4SO4 + 3 H2O
Since ΔG of this reaction is −12.8 kcal/mol, we assign Grel(C2H4SO4) = −12.8 kcal/mol. Because glycolaldehyde has Grel(C2H4O2) = −11.4 kcal/mol, we can quickly see that its bisulfite adduct is 1.4 kcal/mol more stable by comparing the two Grel values. Note that the reaction shown above is to explain the thermodynamic reference states used to calculate Grel and does not represent a prebiotic chemical reaction that actually takes place.
Our results begin with the addition of H2SO3 to form bisulfite analogs of C1 and C2 formose moieties. This is followed by examining the parasitic Cannizzaro reaction where bisulfite adducts reduce its favorability. We then examine how bisulfite analogs in the three steps of the smallest core formose cycle alter the relative kinetics and thermodynamics of the cycle. Finally, we explore the bisulfite analogs of the C2 + C3 → C5 aldol addition where ribose is marginally stabilized over its counterparts. A reminder that all our free energy values are for a model system under neutral conditions at 1 M in implicit solvent, absent catalysts, pH, or concentration changes. In the actual experimental system, these numbers will change. Therefore, while we present actual values of ΔG and ΔG‡, the reader should not interpret these as absolute values corresponding to an experimental system.
3.1. Initial Formation of Bisulfite Adducts and Rearrangements
While H2SO3 can form an adduct with any of the formose compounds, this section discusses the simplest cases: addition to formaldehyde (the C1 food supply) and glycolaldehyde (the linchpin C2 molecule for autocatalysis). Our calculated aqueous free energies establish the thermodynamics and kinetics of adding and rearranging bisulfite moieties; energetics of larger sugars are generally analogous and are discussed in their respective sections.
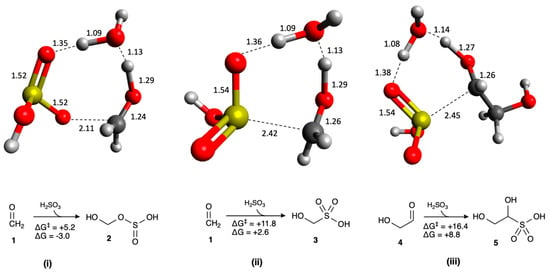
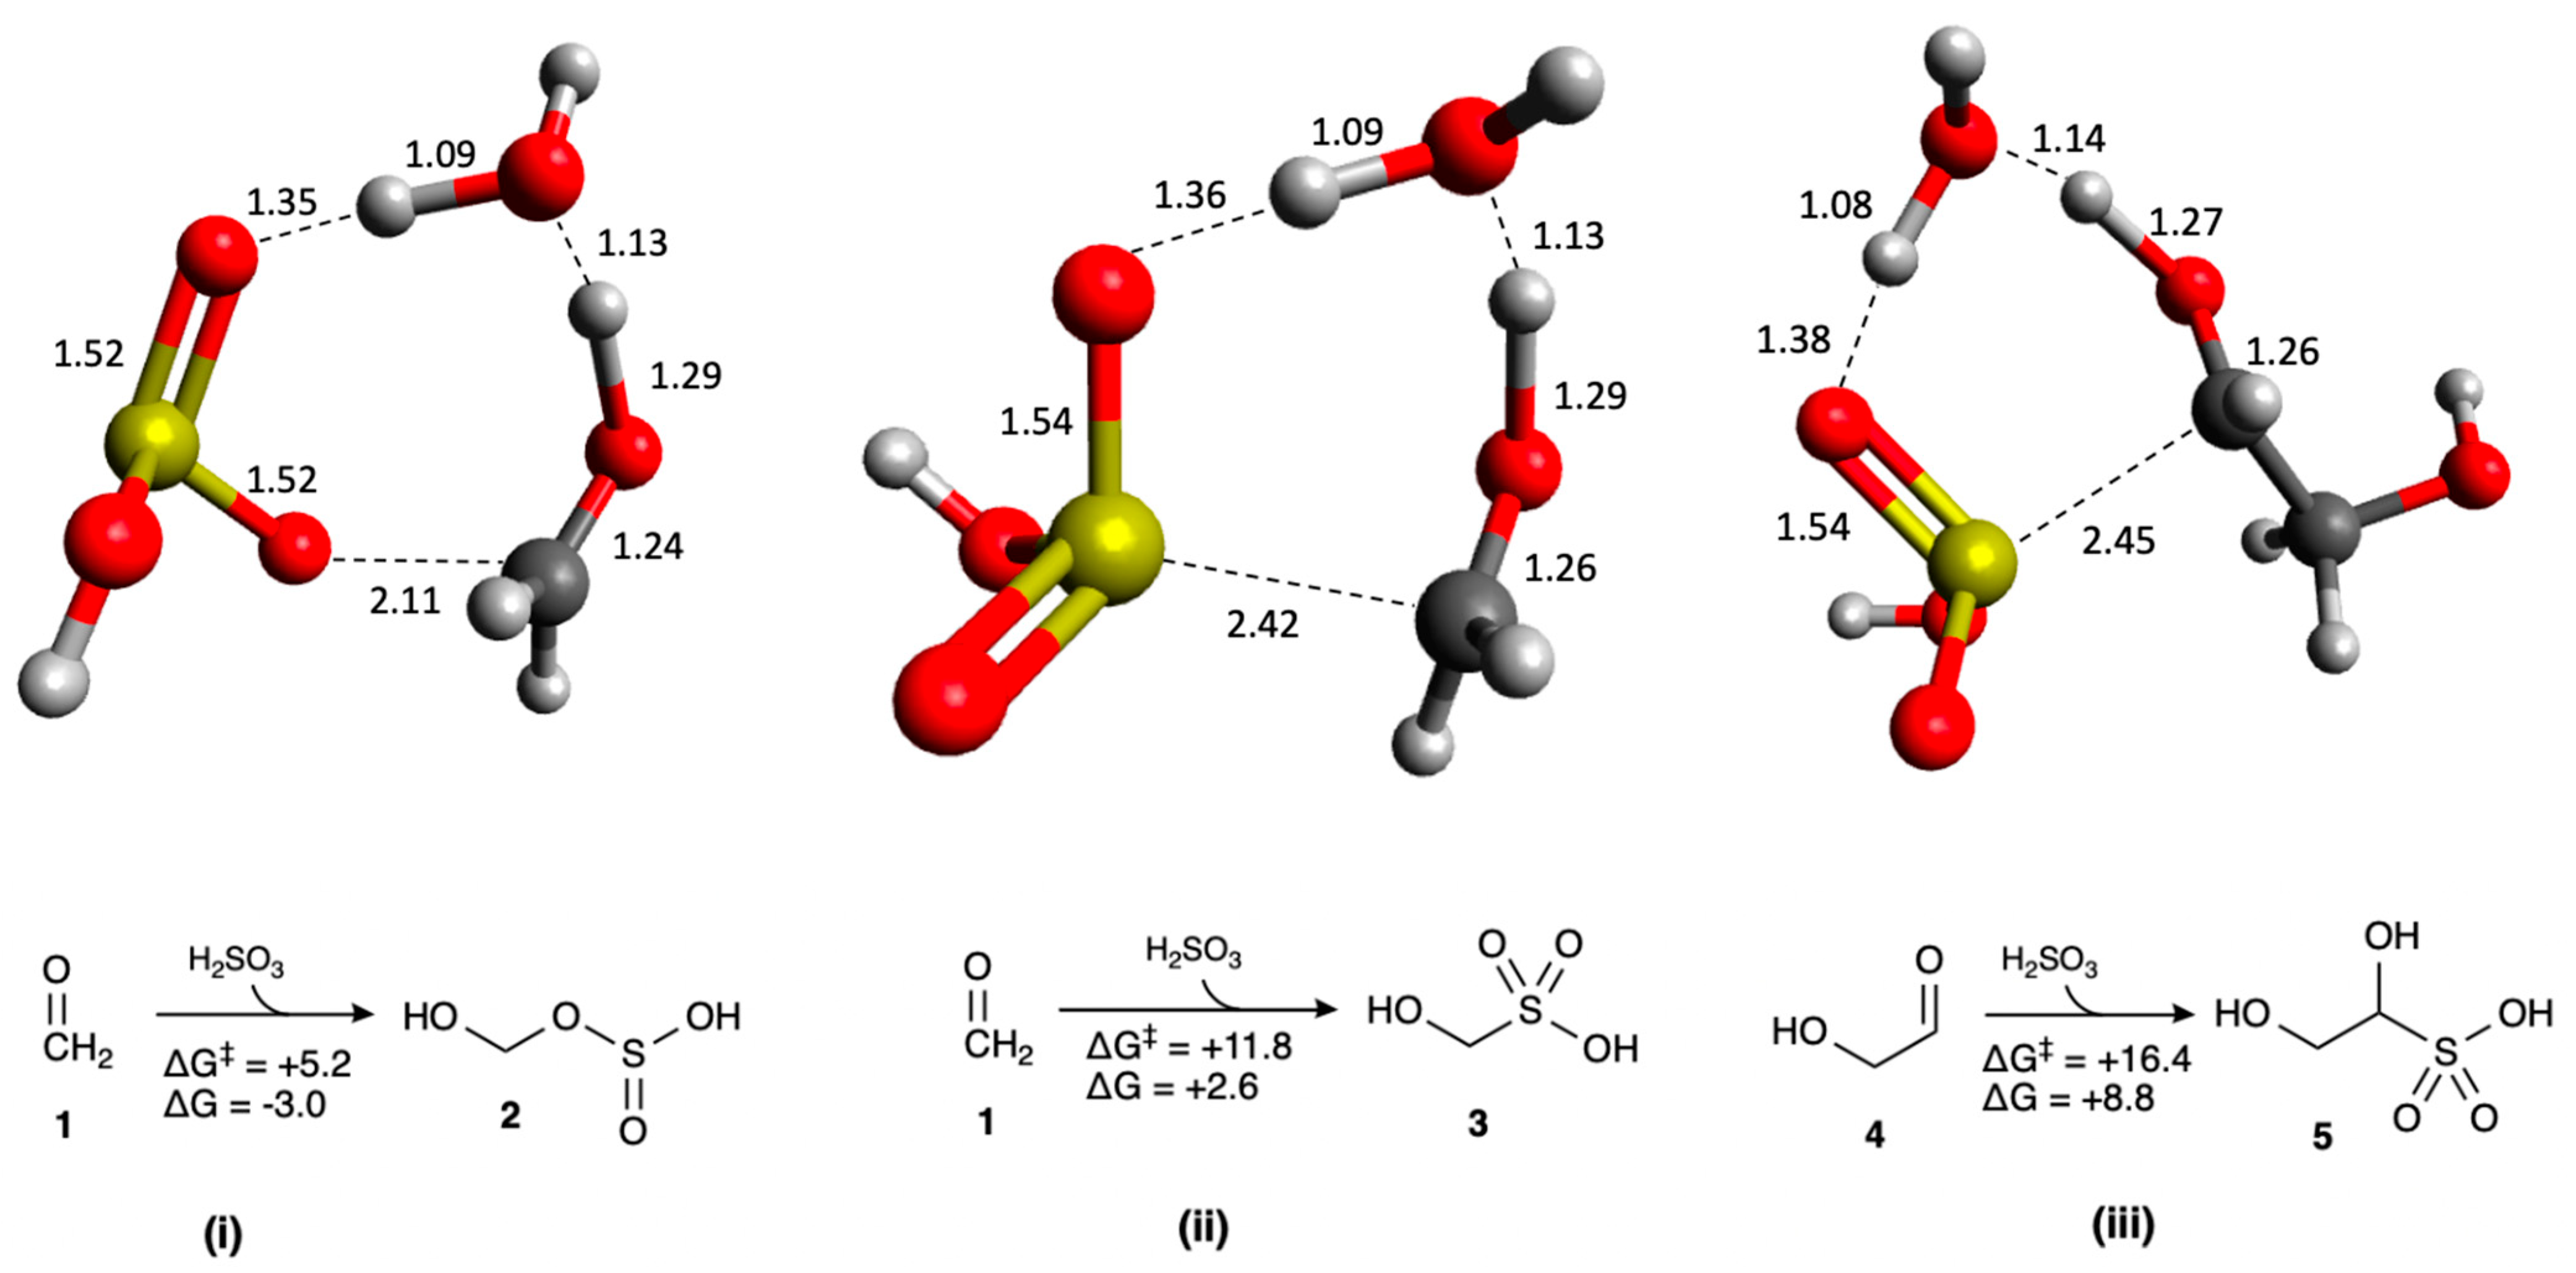
We first consider the addition of H2SO3 to CH2O as shown in Figure 2. The carbonyl carbon of 1 (CH2O) is the electrophilic site. There are two possibilities: If the oxygen of H2SO3 acts as the nucleophile (“O-attack”), the reaction is slightly exergonic in our model (ΔG = −3.0 kcal/mol) with a low barrier (ΔG‡ = +5.2 kcal/mol) and forms the bisulfite adduct 2. Alternatively, if the sulfur of H2SO3 acts as the nucleophile (“S-attack”), formation of the bisulfite adduct 3 is now thermodynamically unfavorable (ΔG = +2.6 kcal/mol) and the barrier has increased (ΔG‡ = +11.8 kcal/mol) although still kinetically feasible. The analogous S-attack on 4 (glycolaldehyde) to form the bisulfite adduct 5, rightmost in Figure 2 is more unfavorable thermodynamically and kinetically. Transition states for all reactions are shown in Figure 2. We expect C3 and C4 S-attack adducts to be similarly unfavorable, and therefore the S-attack molecules will not be considered further in this article; we focus only on O-attack for adduct formation.
Figure 2.
Reactions forming bisulfite adducts: (i) O-attack on CH2O, (ii) S-attack on CH2O, (iii) S-attack on glycolaldehyde (Grel in kcal/mol, bond distances in Å).
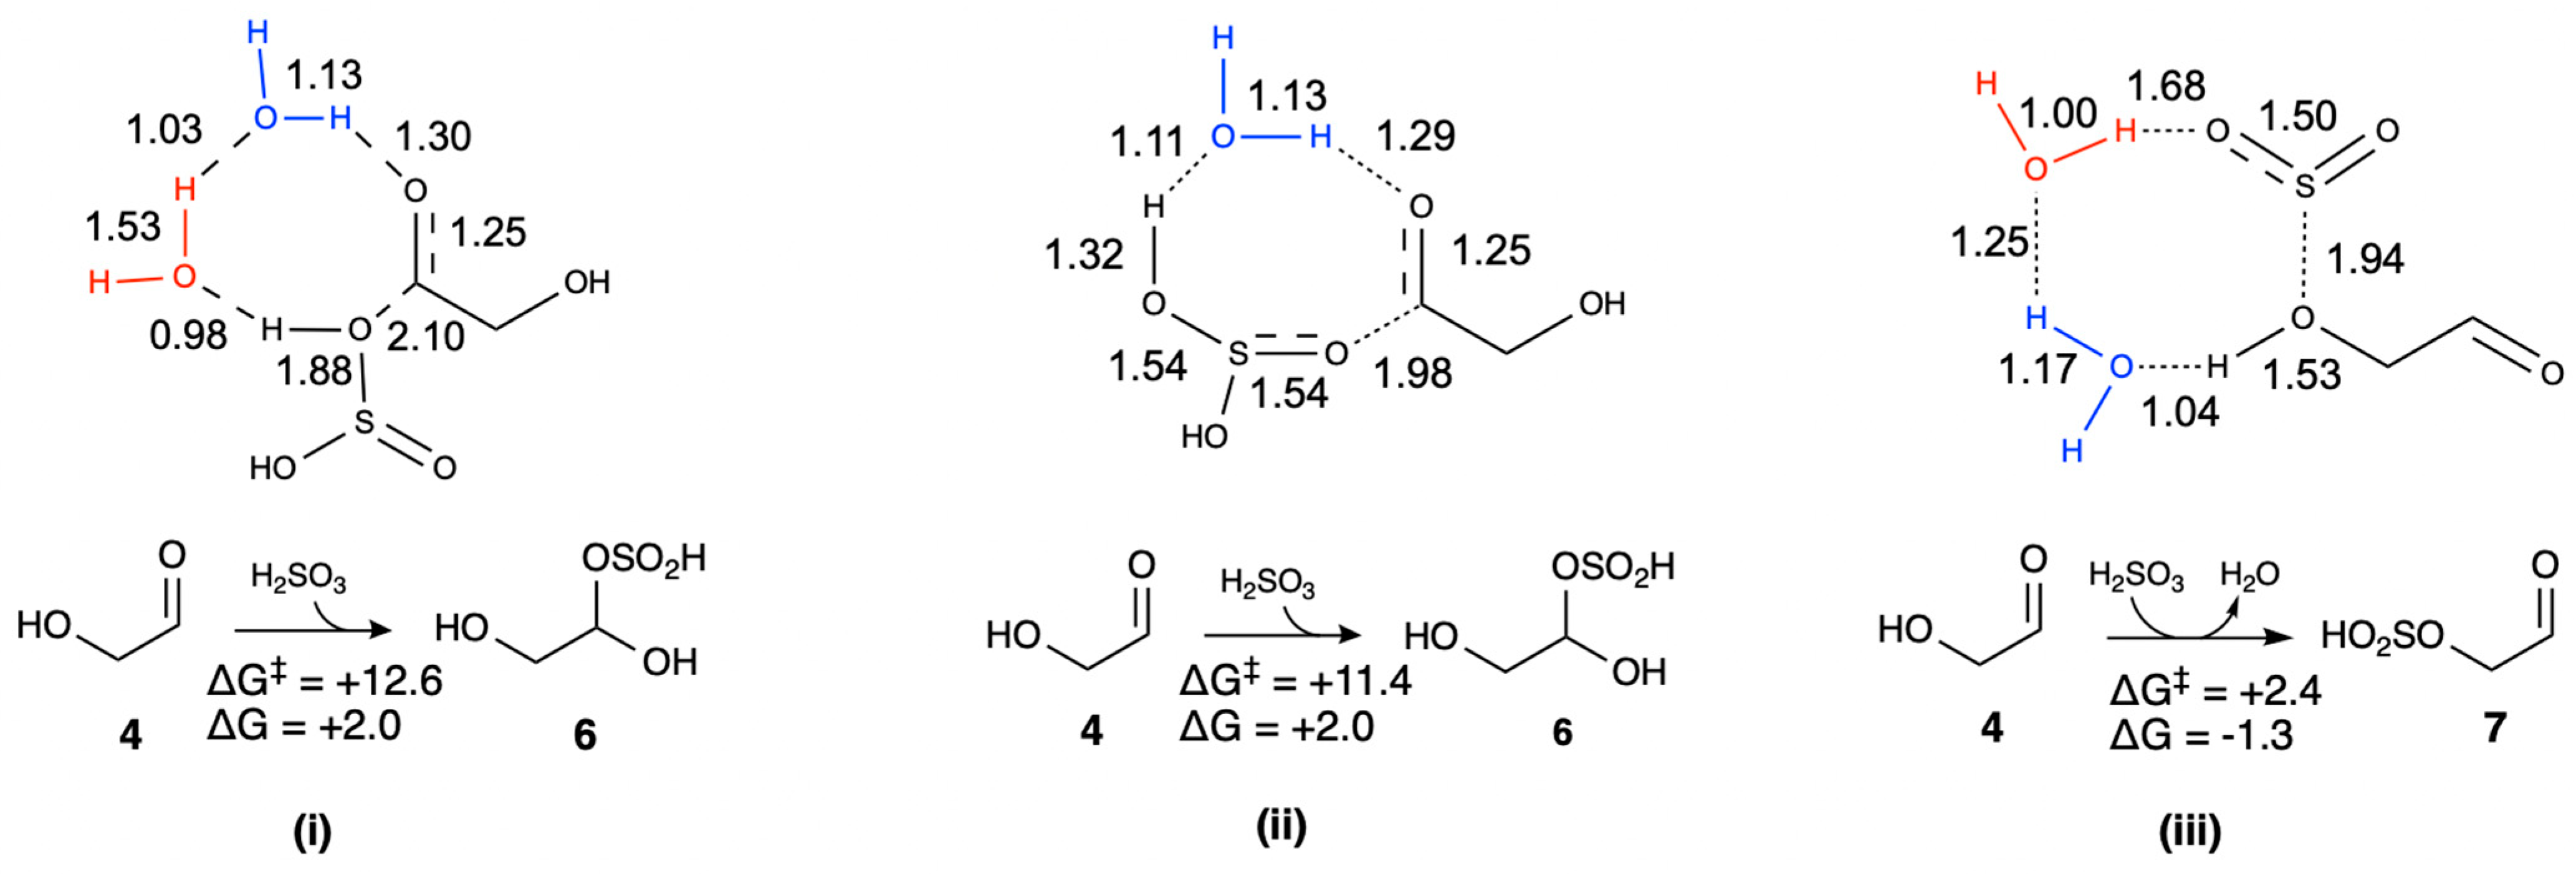
We now consider the addition of H2SO3 and SO2 to 4 in more detail. If bisulfite is the O-attack nucleophile attacking the carbonyl of glycolaldehyde, forming the adduct 6 is thermodynamically uphill (ΔG = +2.0 kcal/mol), although much less so than the S-attack. If the hydroxyl of H2SO3 performs the O-attack (reaction (i) in Figure 3), the barrier is relatively low (ΔG‡ = +12.6 kcal/mol), and the forming O…C bond is 2.10 Å in the transition state. If the sulfoxide (S=O) performs the O-attack (reaction (ii) in Figure 3), the barrier is marginally lower (ΔG‡ = +11.4 kcal/mol), and the forming O…C bond in the transition state is 1.98 Å. The optimal transition state in both cases is an 8-center transition state with relevant transition state distances schematically shown in Figure 3. More detailed transition state geometrical structures are shown in Supplementary Materials. As stated in Computational Methods, we try different numbers and geometries of catalytic water molecules when calculating transition states and only the lowest energy structures are shown.
Figure 3.
Three mechanisms for glycolaldehyde bisulfite adduct formation (Grel in kcal/mol, bond distances in Å).
If instead of reacting at the carbonyl end of glycolaldehyde, the hydroxyl of glycolaldehyde acts as the nucleophile and attacks the sulfur, this yields the bisulfite adduct 7 (reaction (iii) in Figure 3). The optimal transition state prefers H2SO3 to be dehydrated to an SO2 moiety in a 6-center transition state. The forming O…S bond is 1.94 Å. This reaction is marginally exergonic (ΔG = −1.3 kcal/mol) with a low barrier (ΔG‡ = +2.6 kcal/mol). We expect this pathway to be dominant since it is both thermodynamically and kinetically more favorable than the other two pathways.
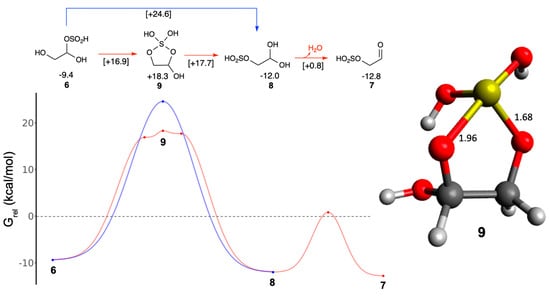
Next, we consider the rearrangement of adduct 6 to adduct 7, which is thermodynamically favorable. It proceeds in two steps: the bisulfite group in 6 moves to the neighboring carbon forming adduct 8 before dehydrating to form 7 as shown in Figure 4. An energy curve is included to orient the reader to our use of Grel values before we get to the complexity of the formose reactions. The Grel of 6 and 8 are −9.4 and −12.0 kcal/mol respectively, therefore ΔG = −2.6 kcal/mol. Since Grel of 7 is −12.8 kcal/mol, the dehydration of 8 to 7 is an additional 0.8 kcal/mol downhill. Thus, the rearrangement of 6 to 7 has overall ΔG = −3.4 kcal/mol. While the energy curve provides a visual aid and is easy to represent in a short linear pathway, such energy diagrams are difficult to represent for complex branching pathways we will examine later in this article. By providing Grel values, the reader can get a quick sense of the global energy landscape.
Figure 4.
Rearrangement of glycolaldehyde bisulfite adduct. (Grel in kcal/mol, bond distances in Å).
For the bisulfite shift between 6 and 8, we envisioned a transient intermediate with the ring structure 9. Since Grel(9) is +18.3 kcal/mol, this intermediate is 18.3 − (−9.4) = 27.7 kcal/mol higher in energy than 6. When we attempted to optimize the transition states on both sides of the intermediate, their energies were very similar (Grel of +16.9 and +17.7 kcal/mol) and marginally below the intermediate. This suggests that our optimized structure 9 is not a stable intermediate and the transition state “hill” may be broad, as can be seen from the three red points on the energy diagram in Figure 4. The ring structure of 9 is asymmetric with S–O bond distances of 1.96 and 1.68 Å respectively. Since the two points on either side of 9 are similar in energy, this suggests that optimizing the transient ring intermediate can be used as a proxy to determine the bisulfite rearrangement barrier without having to optimize the more challenging transition states. We were able to optimize a symmetric transition state with a Grel of +24.6 kcal/mol, but this is 7–8 kcal/mol higher (blue point on the hilltop in Figure 4) than the asymmetric structures. (Structures of all these transition states are in Supplementary Materials.) Regardless, rearrangement via a direct bisulfite shift has a much higher barrier and based on our results we hypothesize that adding and removing the SO2 moiety to be the predominant mechanism since it is kinetically facile with much lower barriers. Actual experimental environmental factor may change the preferred mechanism.
The adduct 7 with an aldehyde group is of similar thermodynamic stability to its hydrate 8 and the hydration barrier is ~13 kcal/mol. These values are very similar to our free energy calculations of glycolaldehyde and its hydrate without bisulfite, which closely matches experimental thermodynamic and kinetic values [35]. Whether 7 or 8 will be more favored will depend on how much water is present. In a dynamic fluctuating environment such as terrestrial hot spring wet-dry cycles proposed by Damer & Deamer [49], the relative abundances of different products will alternate with the cycles and the experimental free energy changes will differ from our calculated values.
3.2. Cannizzaro Reactions of C1 and C2 Bisulfite Adducts
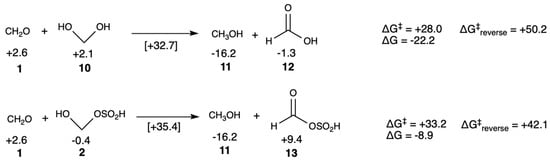
The parasitic Cannizzaro disproportionation is always present in formose systems and detracts from the reactant pool by reducing the concentration of sugars participating in aldol reactions. While the uncatalyzed barrier is relatively high (ΔG‡ = +28.0 kcal/mol) for CH2O reacting with its hydrate 10 to form methanol 11 and formic acid 12 under neutral conditions, the reaction is highly exergonic (ΔG = −22.2 kcal/mol) as shown in Figure 5. Under alkaline conditions, the reaction would be kinetically much more feasible and Cannizzaro products build up [2]. (The Cannizzaro is thermodynamically strongly biased toward products under standard conditions; the reverse reaction is very endergonic and the reverse barrier would still be high even with a catalyst.)
Figure 5.
Cannizzaro reactions of formaldehyde and its bisulfite adduct (Grel in kcal/mol).
In the previous section, we saw that bisulfite adds to formaldehyde to form adduct 2. The Cannizzaro reaction of CH2O and 2 to form methanol and the corresponding bisulfite ester 13 is even more kinetically unfavorable under neutral or acidic conditions, and much less exergonic (ΔG = −8.9 kcal/mol). (See Supplementary Materials for the transition state structure.) While the barrier will be lowered in alkaline conditions the barrier with bisulfite is ~5 kcal/mol higher than without bisulfite. Thus, we expect that the presence of H2SO3 suppresses the Cannizzaro reaction. This was observed experimentally by Kawai et al. [32]; our results support their argument that adduct 2 provides a mechanism to sequester formaldehyde and suppress the likelihood of Cannizzaro reactions, but does not eliminate it entirely. However, sequestering formaldehyde will also slow down aldol additions involving CH2O.
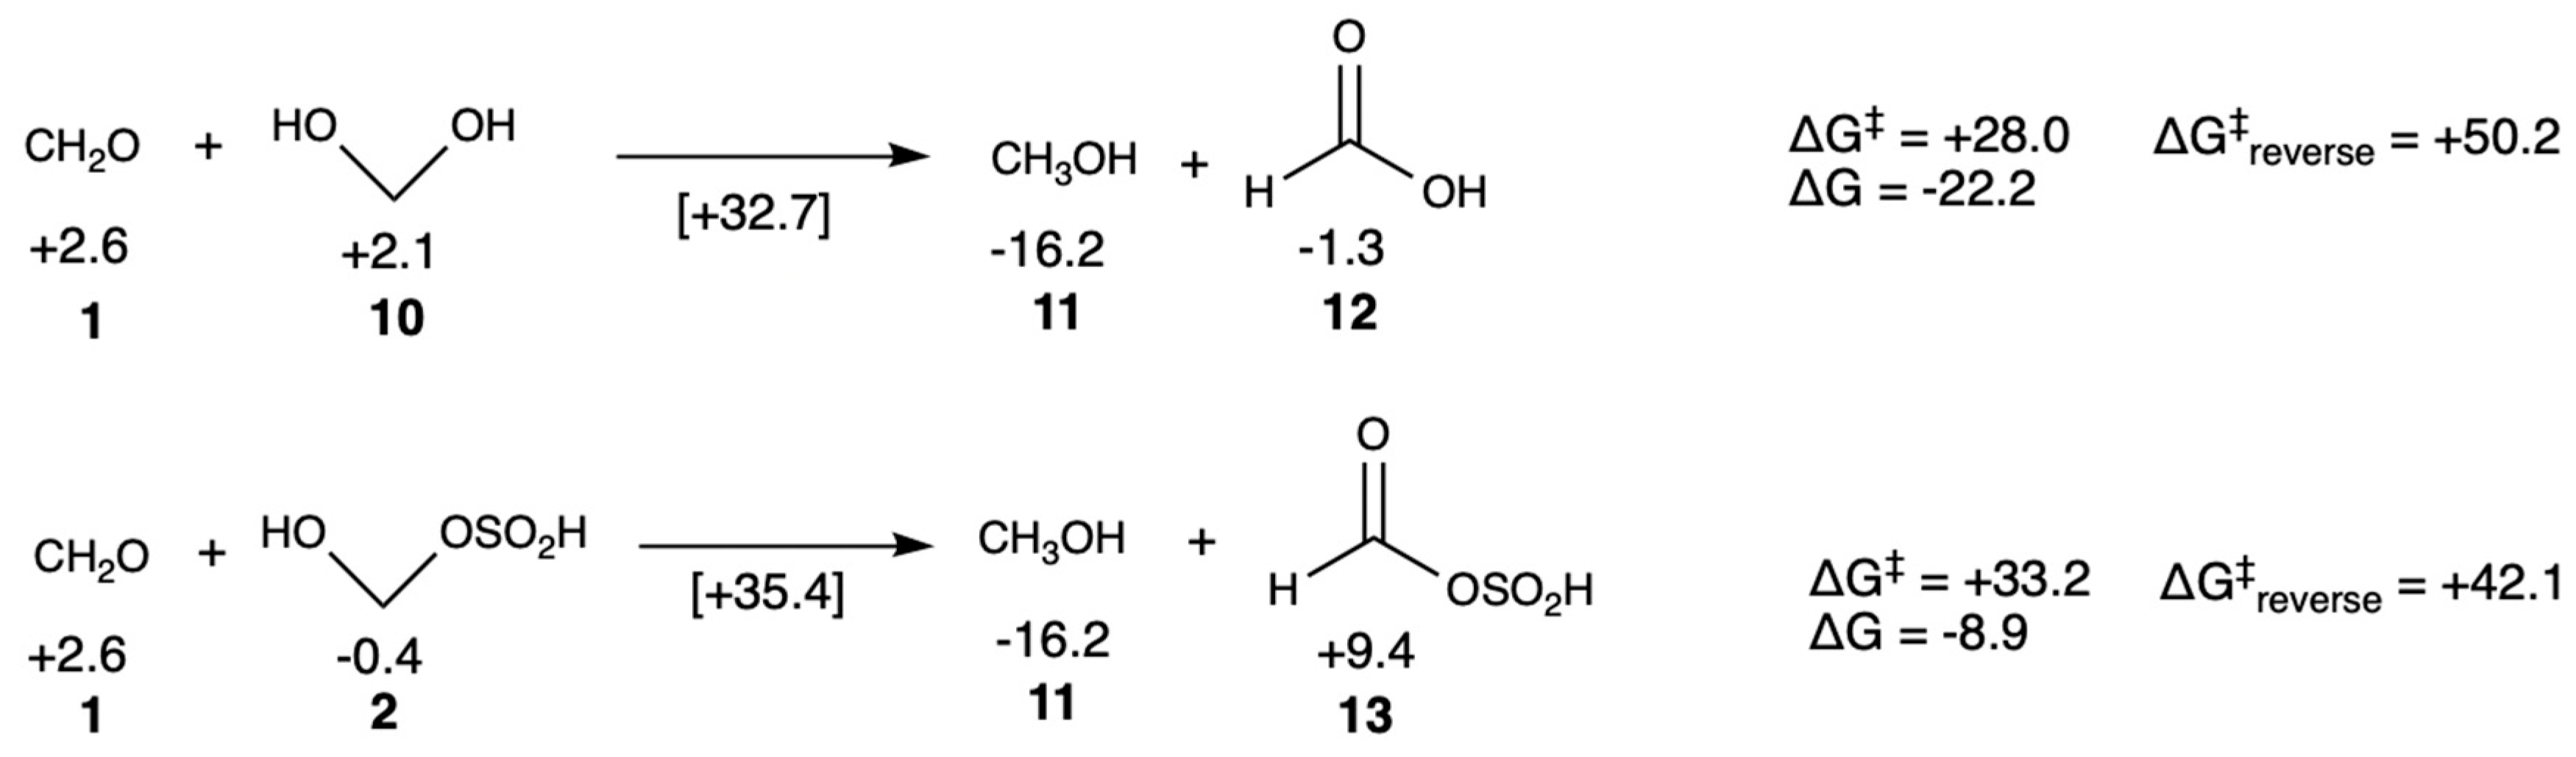
Although glycolaldehyde can self-disproportionate, its low concentration compared to formaldehyde in the early stages of the formose cycle makes this less likely. On the other hand, cross-Cannizzaro reactions between C1 and C2 aldehydes might occur. The results are shown in in Figure 6. For reaction (i) with no sulfur analog, ΔG = −16.3 kcal/mol and ΔG‡ = +32.7 kcal/mol under neutral conditions. If the bisulfite group is on glycolaldehyde and does not participate, as shown in reaction (ii), ΔG is not too different although the barrier is lowered by ~5 kcal/mol. If the bisulfite group is on formaldehyde, the reaction is now much less exergonic, as shown in reactions (iii) and (iv), with ΔG values of −3.1 and −5.2 kcal/mol respectively, as expected from our results in the previous section. This is because the favorable reduction of the C2 compound (forming 14 and 15) is largely compensated by the unfavorable oxidation of 2 to 13; the electron withdrawing bisulfite group disfavors its further oxidation in our computational model.
Figure 6.
Cross-Cannizzaro reactions of glycolaldehyde and the bisulfite adduct of formaldehyde (Grel in kcal/mol).
3.3. Glycolaldehyde-Bisulfite Reactions: Enolization, Aldol Addition and SO2 Elimination
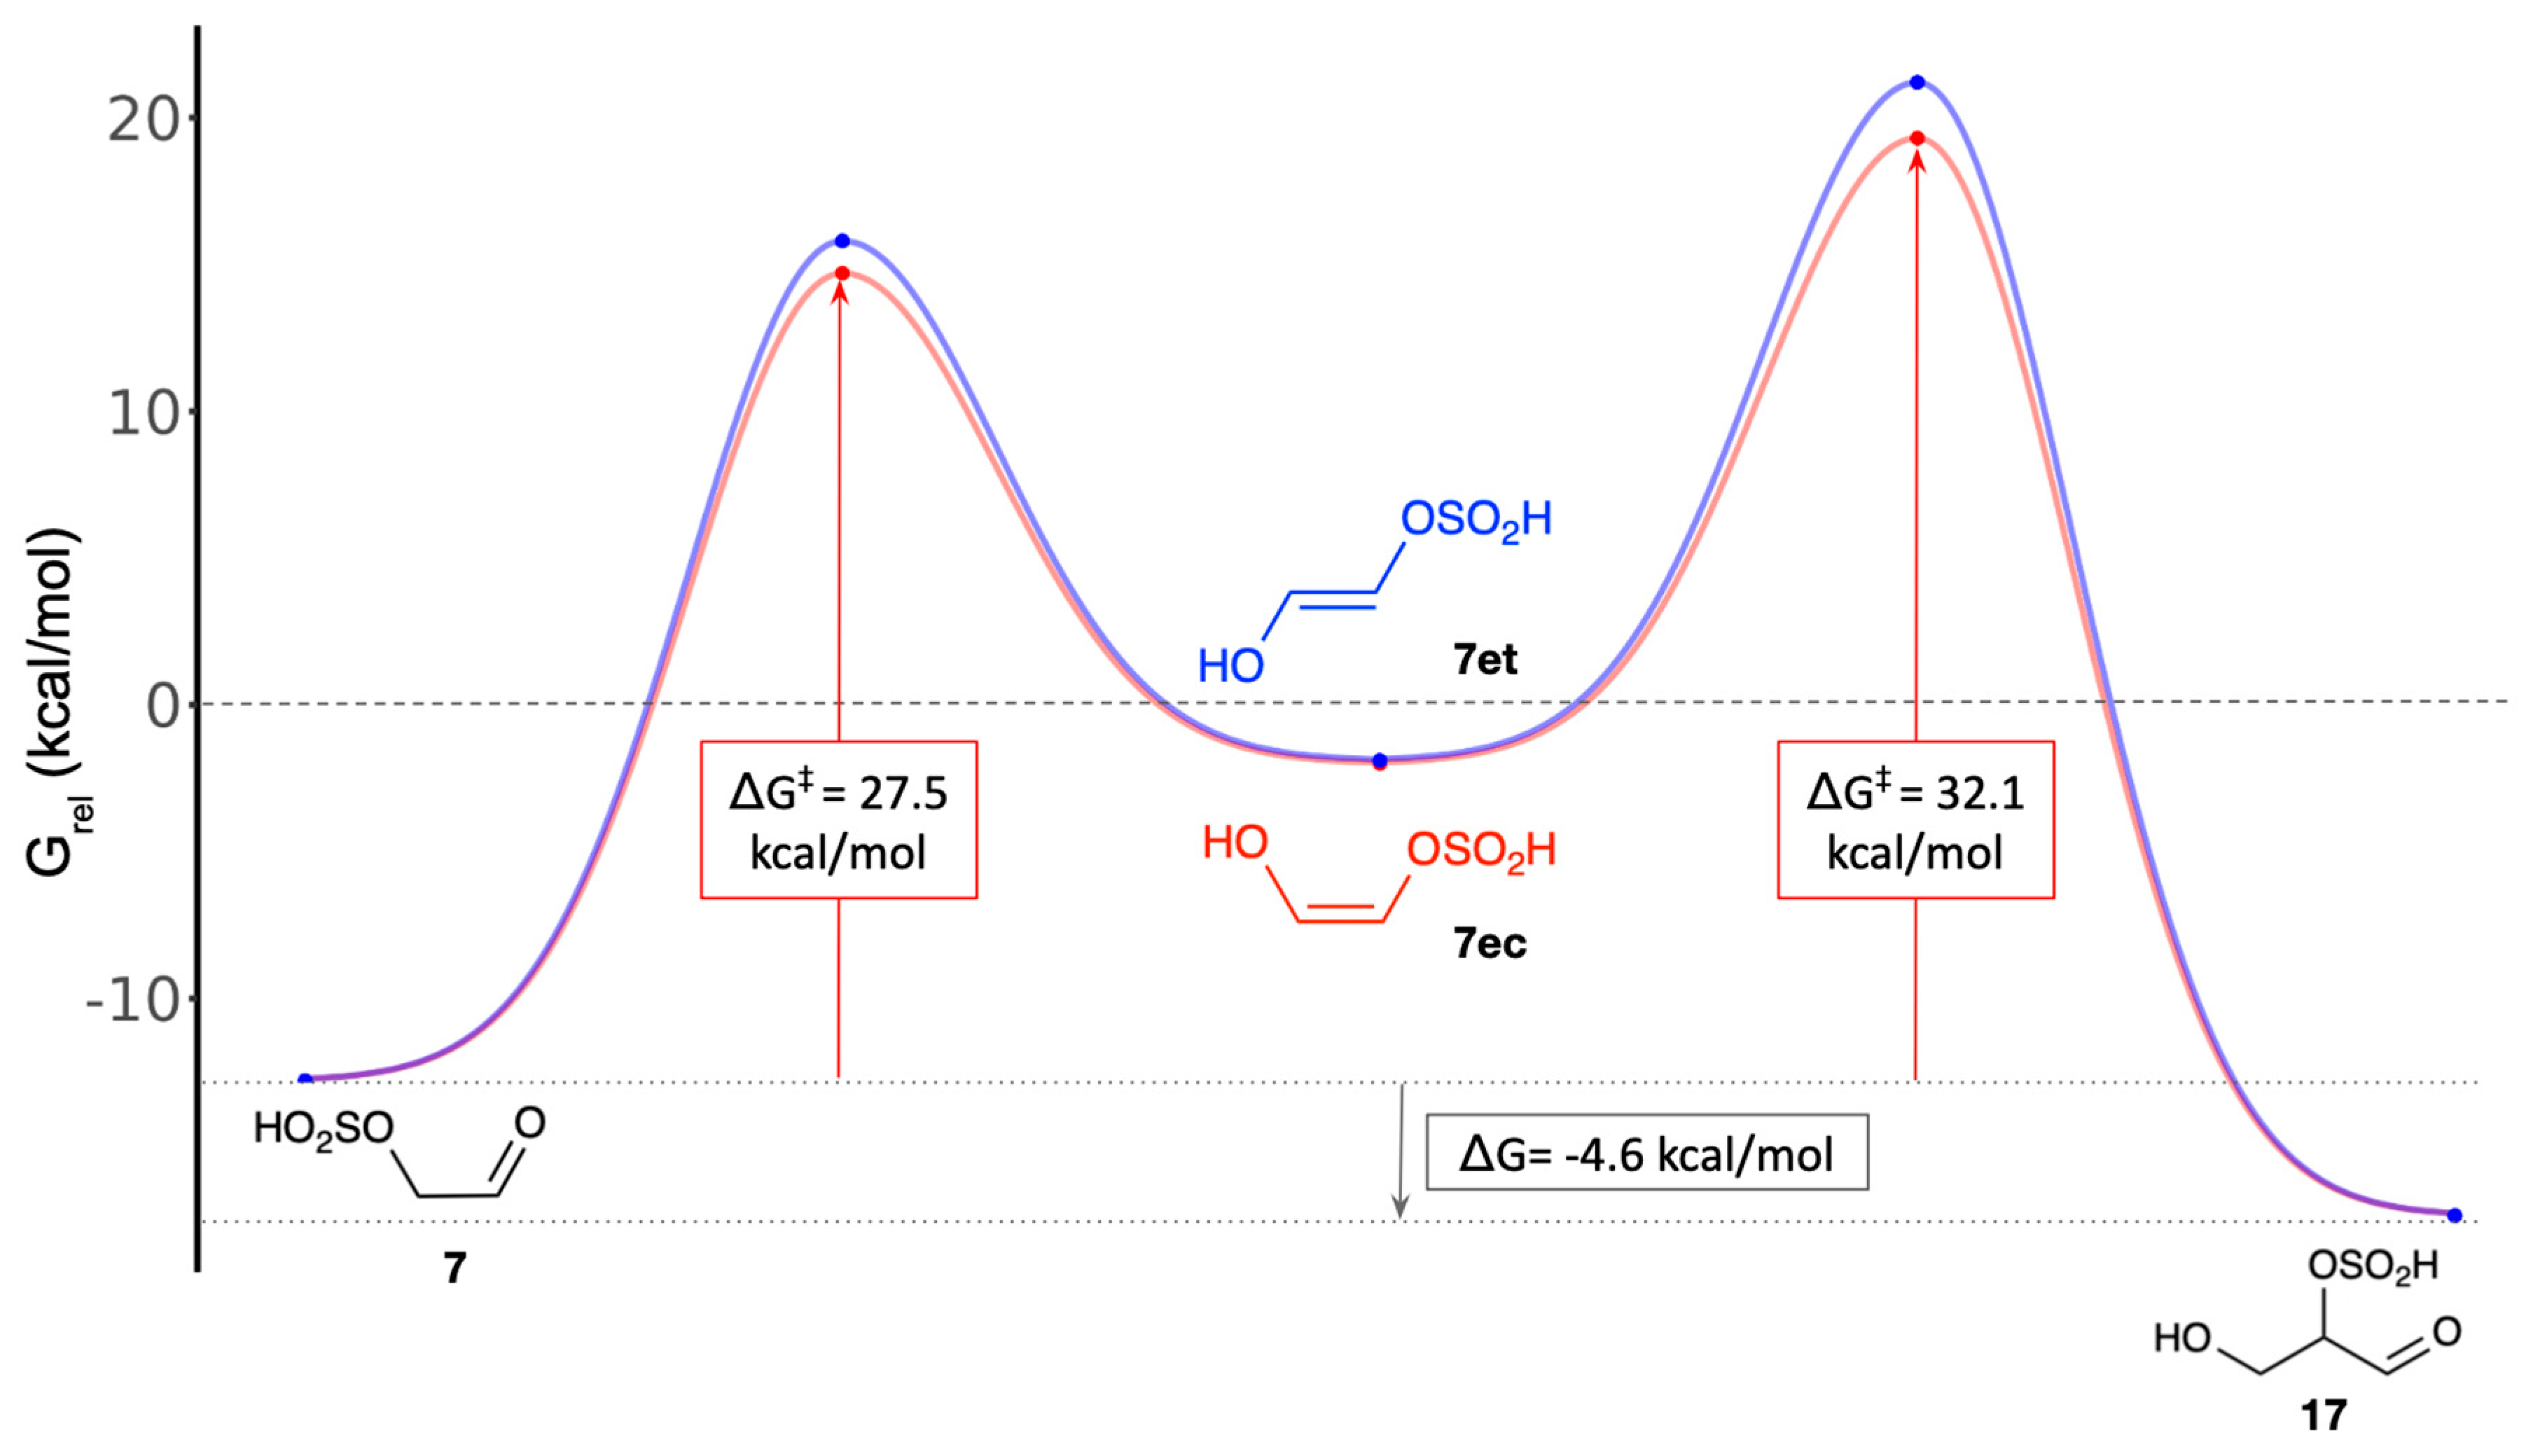
As discussed in Section 3.1, the most stable glycolaldehyde bisulfite adduct is 7, which is favorably generated by the addition of H2SO3 to glycolaldehyde. To participate in aldol reactions in the formose system, it must first enolize. The barrier to enolize under neutral conditions is relatively high (ΔG‡ = +27.5 kcal/mol), although under alkaline formose conditions, we hypothesize that this barrier would be substantially lowered. Forming the cis-enol is favored over the trans-enol both kinetically and thermodynamically, as shown in Figure 7. (7ec is 0.1 kcal/mol lower in energy than 7et, benefiting from intramolecular hydrogen bonding.) An enediol is now susceptible to aldol addition with CH2O acting as the electrophile [50]. In the non-sulfur analog, the C2 + C1 aldol addition would form glyceraldehyde 16; if the bisulfite group was retained throughout, the glyceraldehyde bisulfite adduct 17 is formed. As shown in Figure 7, the overall aldol addition is exergonic (ΔG = −4.6 kcal/mol) but the uncatalyzed barrier is high (ΔG‡ = +32.1 kcal/mol from the aldehydic reactants). In neutral aqueous solution in the absence of catalysts, the reaction is unlikely to occur.
Figure 7.
Enolization and C2 + C1 aldol addition of the bisulfite adduct.
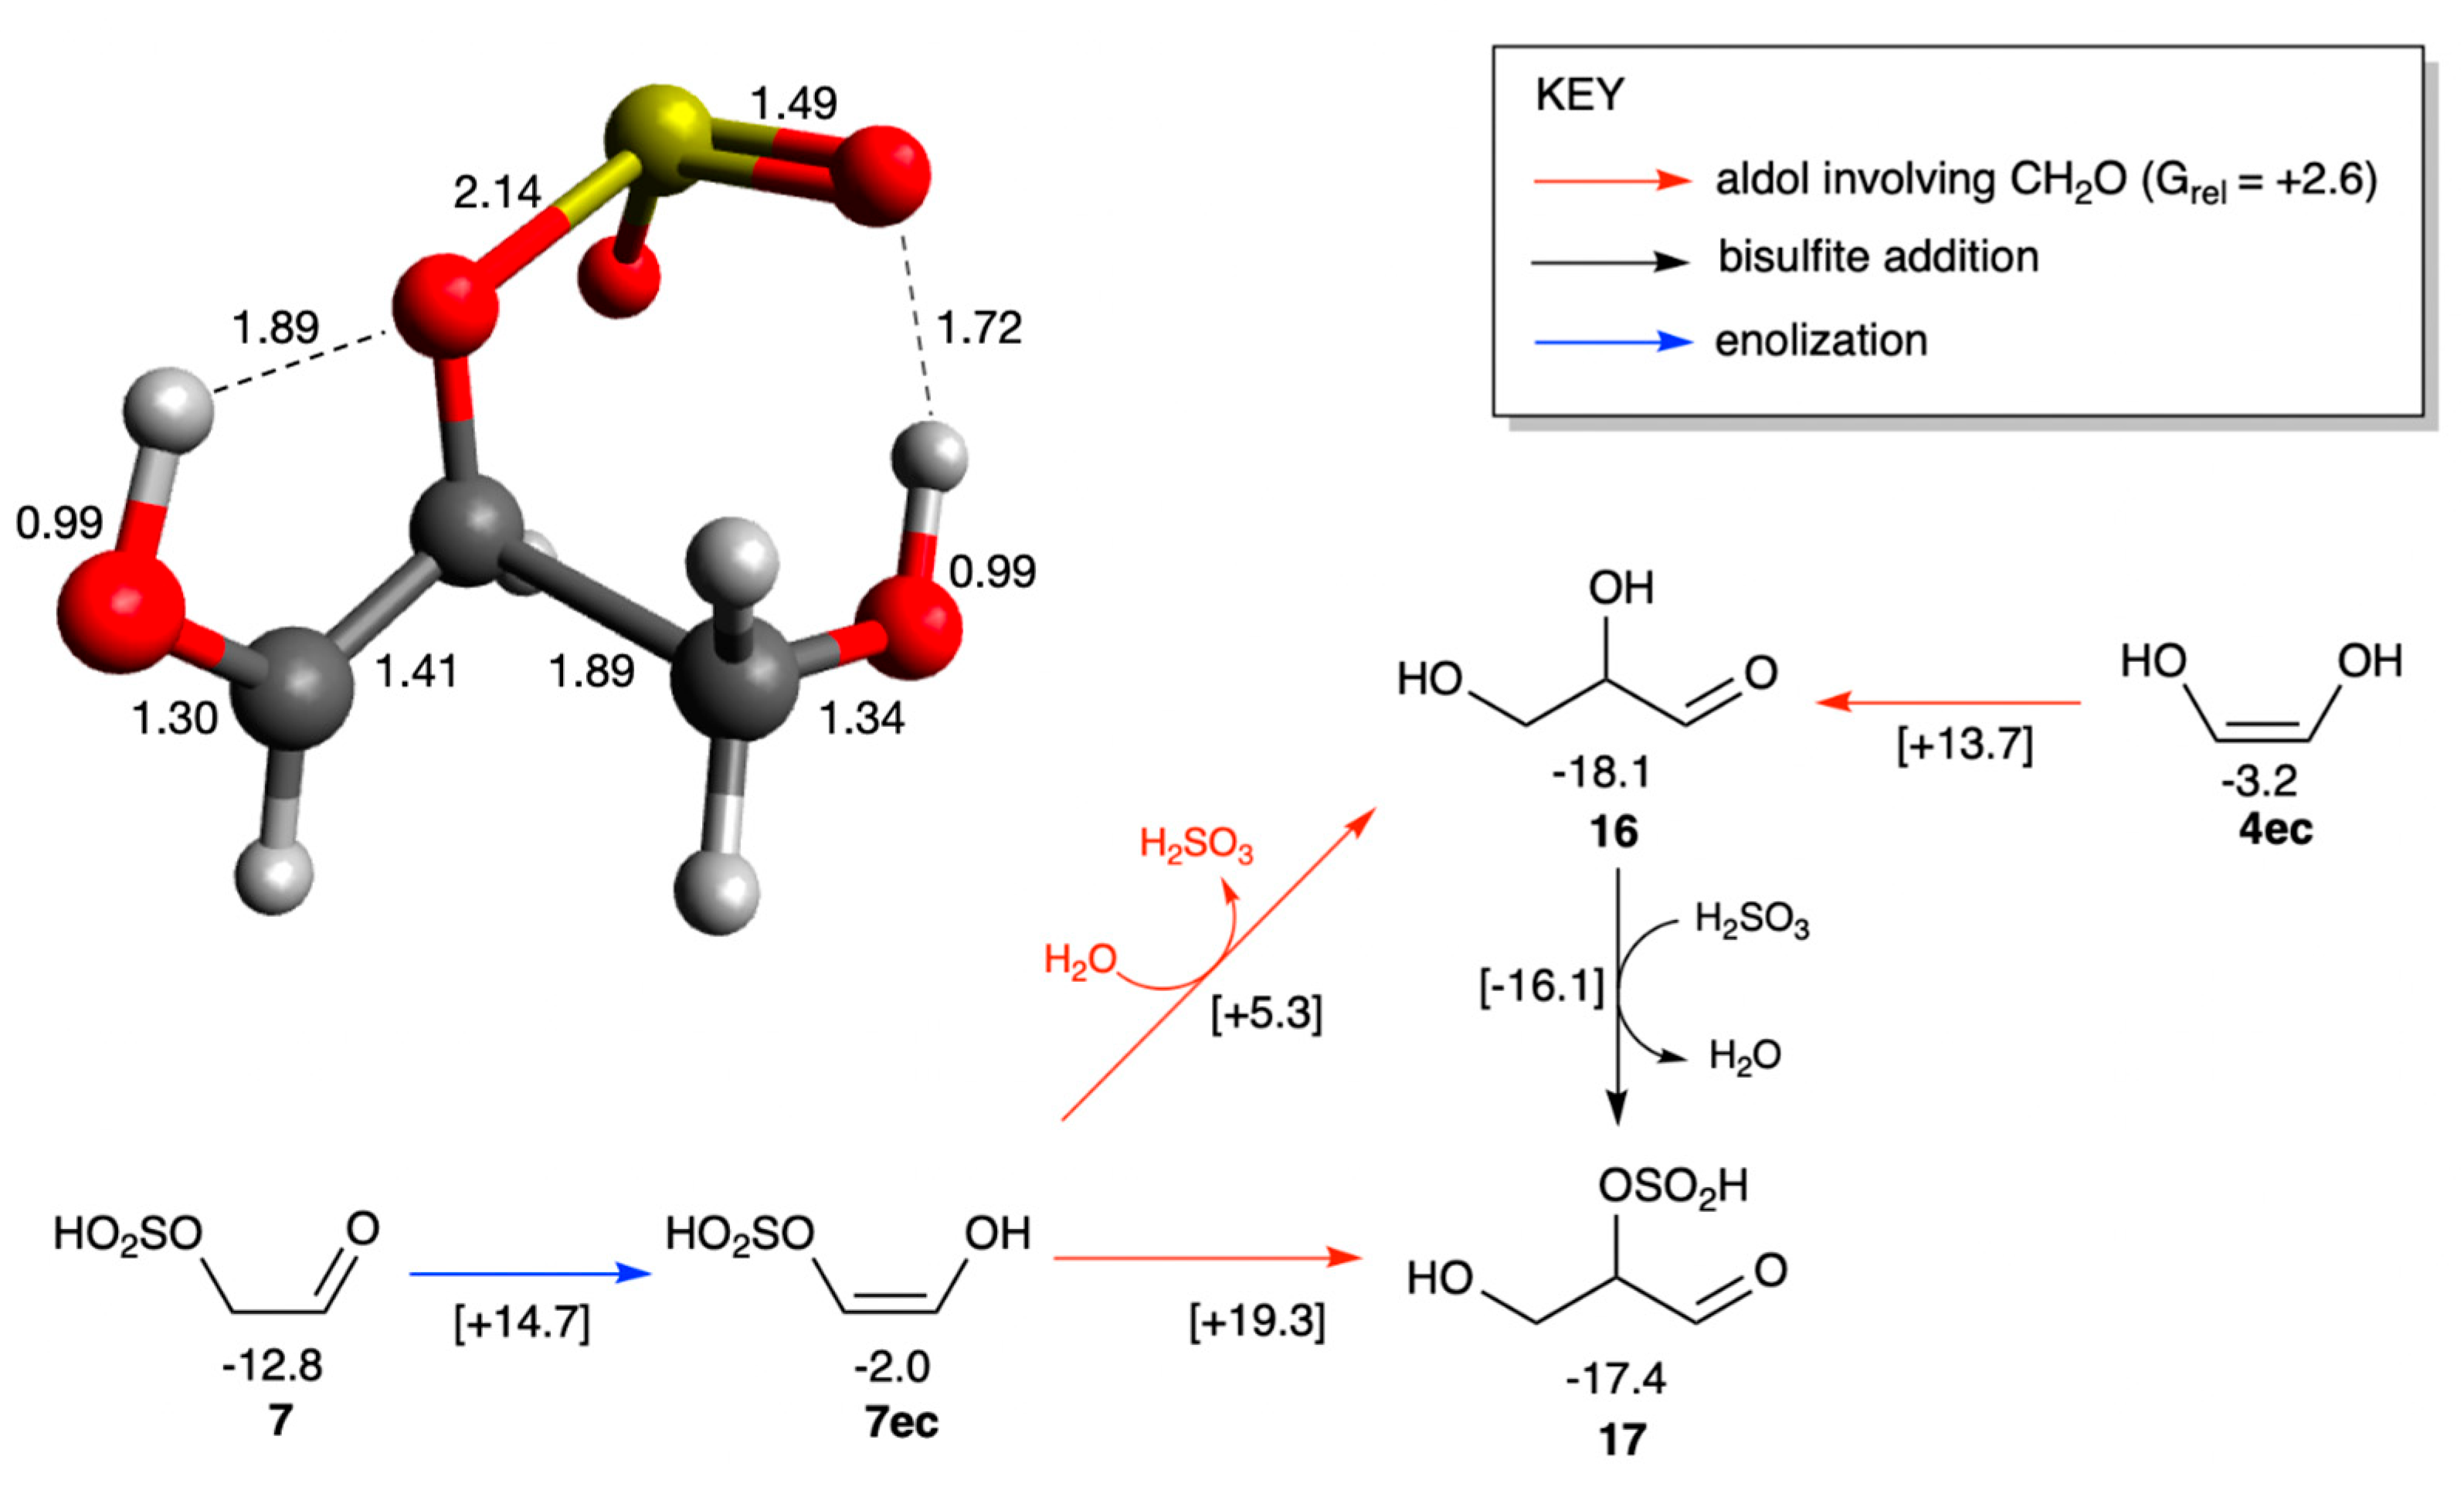
The bisulfite moiety however provides an opportunity for intramolecular catalysis. When we tried to locate a transition state whereby the bisulfite acts as both proton donor and acceptor to facilitate the aldol addition, we always observed the elimination of SO2 concomitant with the aldol addition as shown in Figure 8 by the transformation of 7ec + 1 to form 16. The breaking O…S bond is 2.15 Å in the transition state, shown schematically in Figure 8 (see Supplementary Materials for transition state figures of both enolization and aldol addition steps). With intramolecular catalysis (and SO2 elimination), the barrier for aldol addition is reduced by 14 kcal/mol compared to 7ec + 1 → 17 (the difference in Grel of the two transition states with values of +5.3 and +19.3 kcal/mol). In comparison to the bisulfite-free analog, 4ec + 1 → 16, intramolecular catalysis lowers the barrier by ~8 kcal/mol. Because only a limited number of explicit solvent molecules are included in our transition states, these estimations are qualitative and in an actual system, SO2 elimination will depend on the pH and free energies of hydration.
Figure 8.
Competing aldol additions of the bisulfite enediol adduct (Grel in kcal/mol, bond distances in Å).
As expected, addition of H2SO3 to glyceraldehyde (16) forming the adduct 17 is marginally endergonic (ΔG = +0.7 kcal/mol), not too different from what we found in the C2 case. The ~2 kcal/mol barrier is very low, thus adding and removing the bisulfite group is kinetically feasible and we expect additions and removals to be constantly taking place in a reaction mixture. Our findings suggest that the bisulfite moiety may play a potentially catalytic role in aldol reactions, at least in our model system.
3.4. Glyceraldehyde Bisulfite Reactions: Isomerizations and Aldol Additions
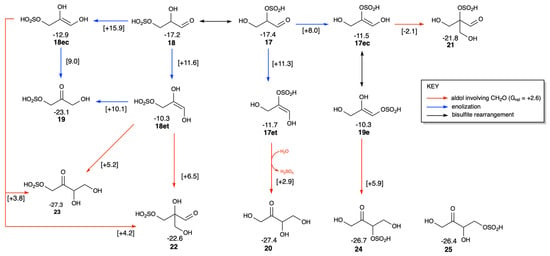
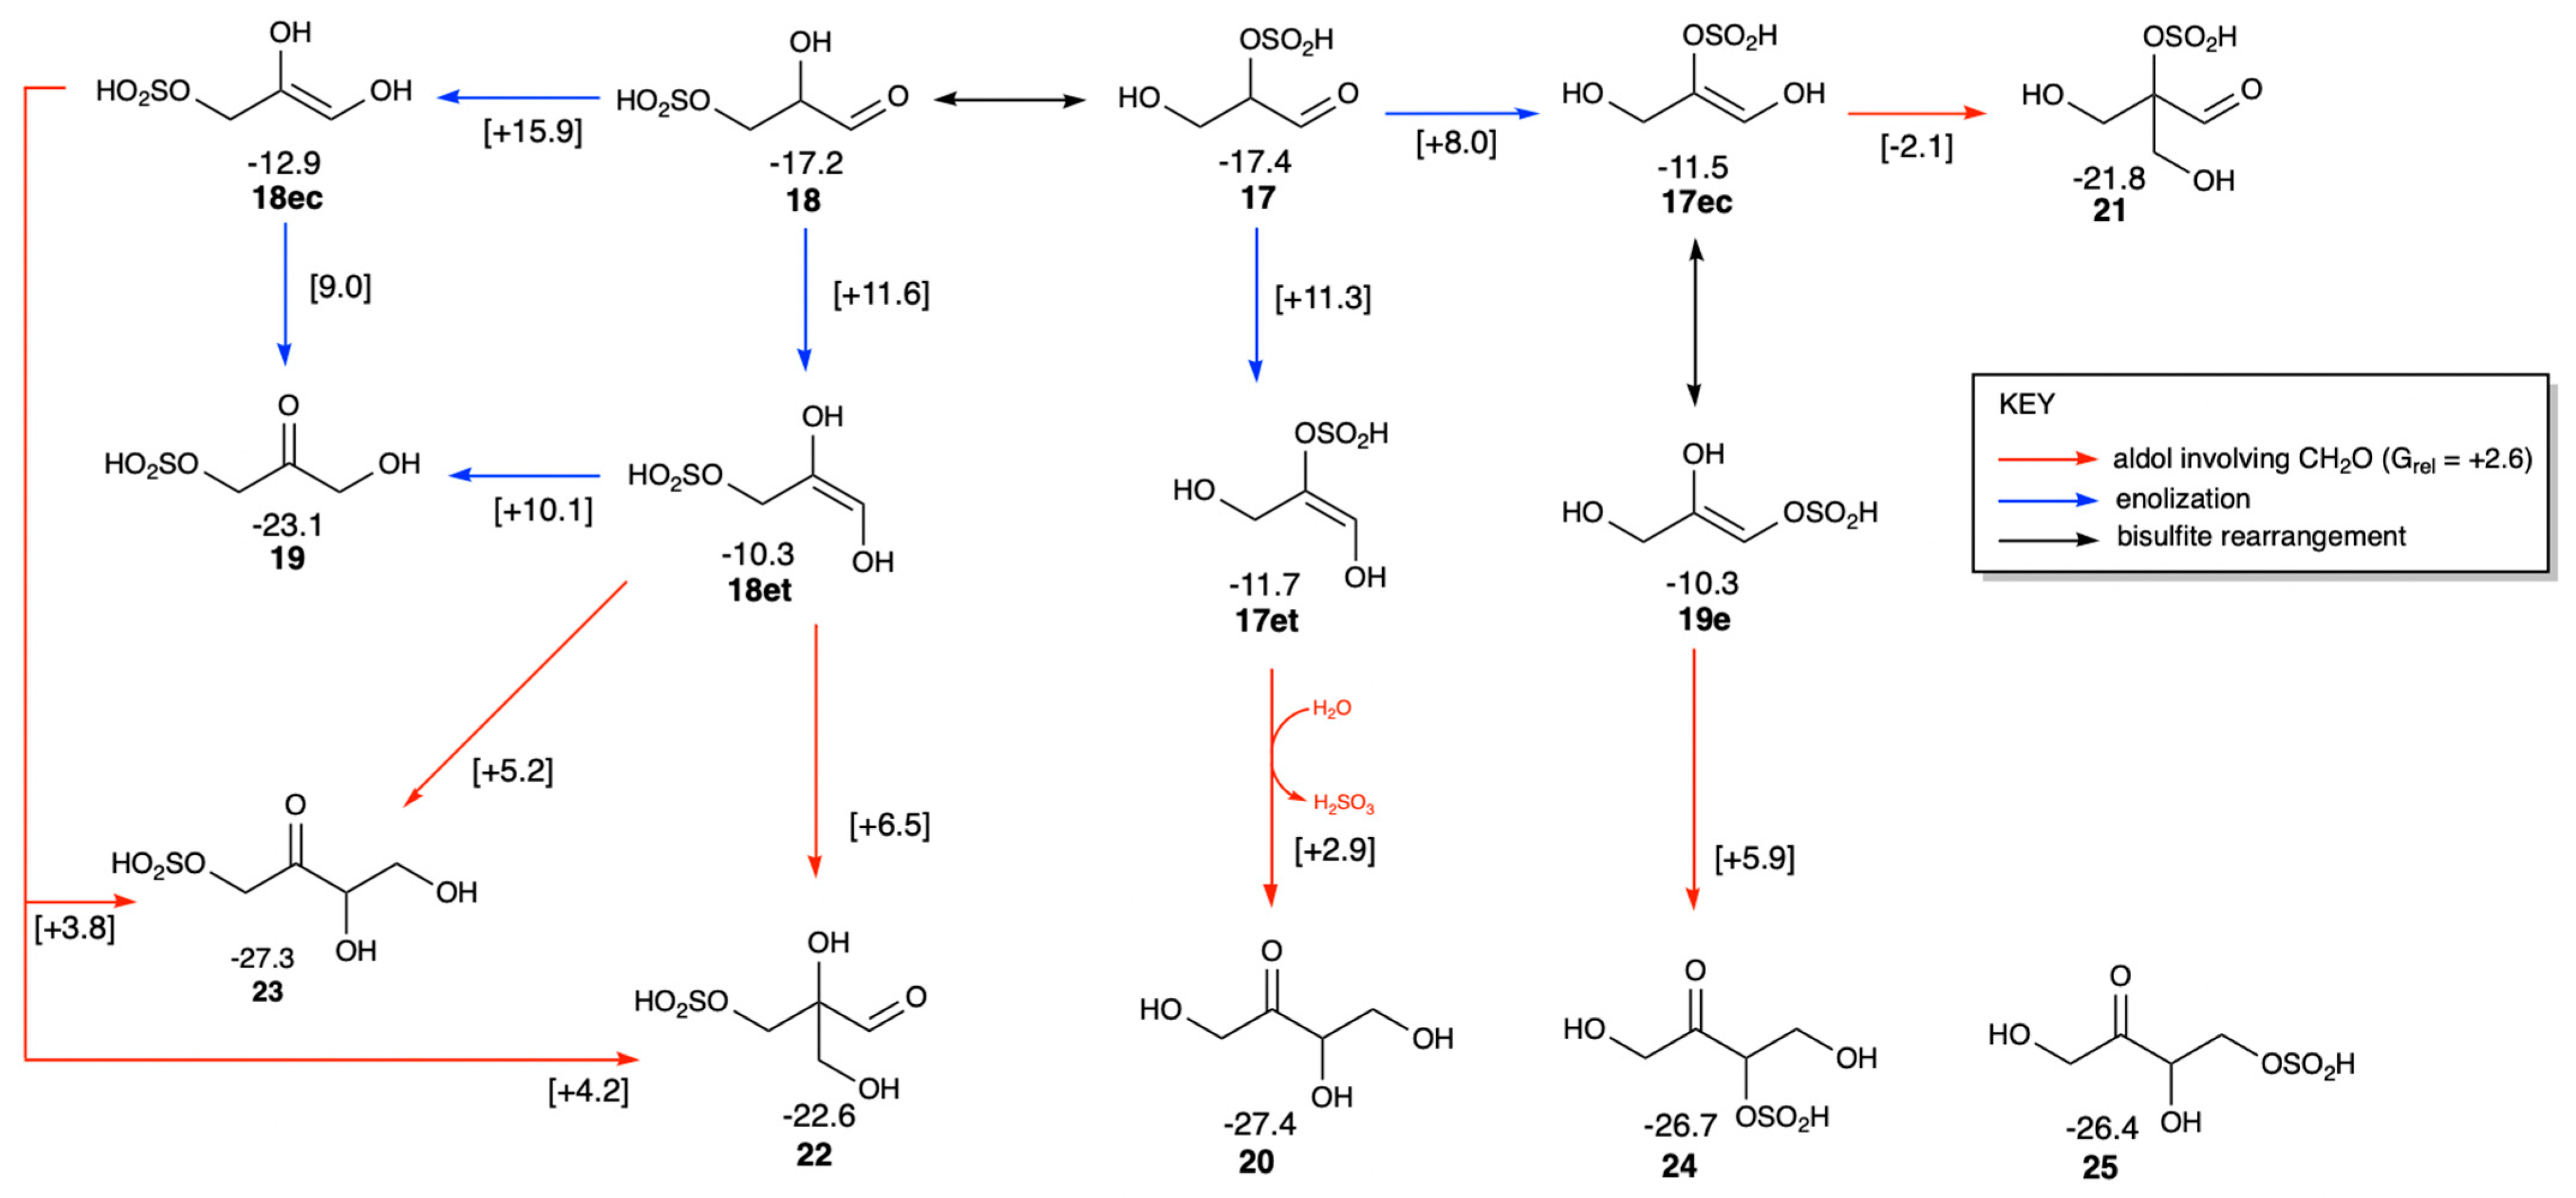
Since the bisulfite moiety can be easily added and removed, alongside adduct 17 we expect to also observe adduct 18 with bisulfite on the terminal rather than central carbon, as shown in Figure 9. They have similar stability with Grel values of −17.4 and −17.2 kcal/mol respectively. In our model, the most likely mechanism for bisulfite rearrangement is removing SO2 in one position and adding it to the other. In previous sections, we saw that this route is kinetically much more feasible with very low barriers. Both aldehydes can enolize forming 17ec, 17et, 18ec and 18et. Not surprisingly, the thermodynamic sink of the trioses is dihydroxyacetone and its adduct 19 (enediol counterpart 19e). In Figure 9, enolizations among C3 species have blue arrows, while bisulfite rearrangements have black arrows.
Figure 9.
Isomerization and aldol addition of glyceraldehyde bisulfite adducts (Grel in kcal/mol).
The C3 + C1 → C4 aldol additions in Figure 9 have red arrows. The most feasible is the intramolecularly catalyzed addition converting 17 to 20 (erythrulose), concomitant with SO2 removal. The overall reaction 17 + 1 + H2O → 20 + H2SO3 is exergonic with ΔG = −27.4 − (−17.4 + 2.6) = −12.6 kcal/mol; erythrulose is the thermodynamic sink of the tetroses. Similar to the C2 + C1 → C3 aldol addition discussed in the previous section, the presence of bisulfite lowers the aldol addition barrier substantially compared to its non-sulfur analog.) We were only able to find a transition state for the second step for the trans-enediol adduct 17et, while its cis counterpart proceeded to form the branched aldose 21.
While kinetically more favorable, the reaction 17 + 1 → 21 is much less exergonic with ΔG = −21.8 − (−17.4 + 2.6) = −7.3 kcal/mol, and is a dead-end reaction which cannot proceed to downstream products because 21 is non-enolizable. This non-productive reaction is therefore parasitic to the formose cycle, but will depend on the availability of enolizable species and the actual kinetics in aqueous solution. Its transition state (see Supplementary Materials) shows that bisulfite acts as an intramolecular proton shuttle, facilitating a lower barrier reaction. Similarly, 18 can form its corresponding branched aldose 22 (less sterically hindered than 21 and therefore marginally more stable by 0.8 kcal/mol). Since the bisulfite moiety is far from the active bond forming and breaking in the transition state and unable to participate in catalysis, its barrier is correspondingly higher. The reaction 18 + 1 → 22 is also parasitic to the formose cycle.
If the bisulfite adduct is not involved (and SO2 is not eliminated) in forming C4 ketose products, there are two more possibilities: 18 + 1 → 23 and 17 + 1 → 24. The erythrulose adducts 23 and 24 have bisulfite at carbons 1 and 3 respectively. Both reactions are similarly exergonic (ΔG = −12.7 and −11.9 kcal/mol respectively) although they have higher barriers than the intramolecular catalyzed 17 + 1 + H2O → 20 + H2SO3 reaction discussed earlier. While there is no direct aldol addition route to ketose adduct 25 with bisulfite at carbon 4, removal and addition of SO2 should be facile. Since the Grel values of 20, 23, 24, and 25 are very similar, computationally this suggests a mixture of all these C4 ketoses may form at equilibrium; however kinetic factors and environmental conditions could skew the distribution. Of the three adducts, 23 (with bisulfite on carbon 1) is marginally more stable because of favorable intramolecular hydrogen bonding with the carbonyl.
3.5. Retro-Aldol Splitting of C4 Tetroses
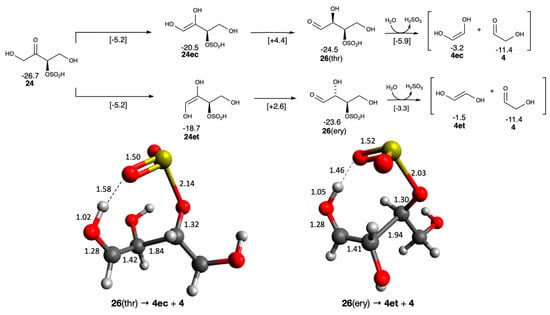
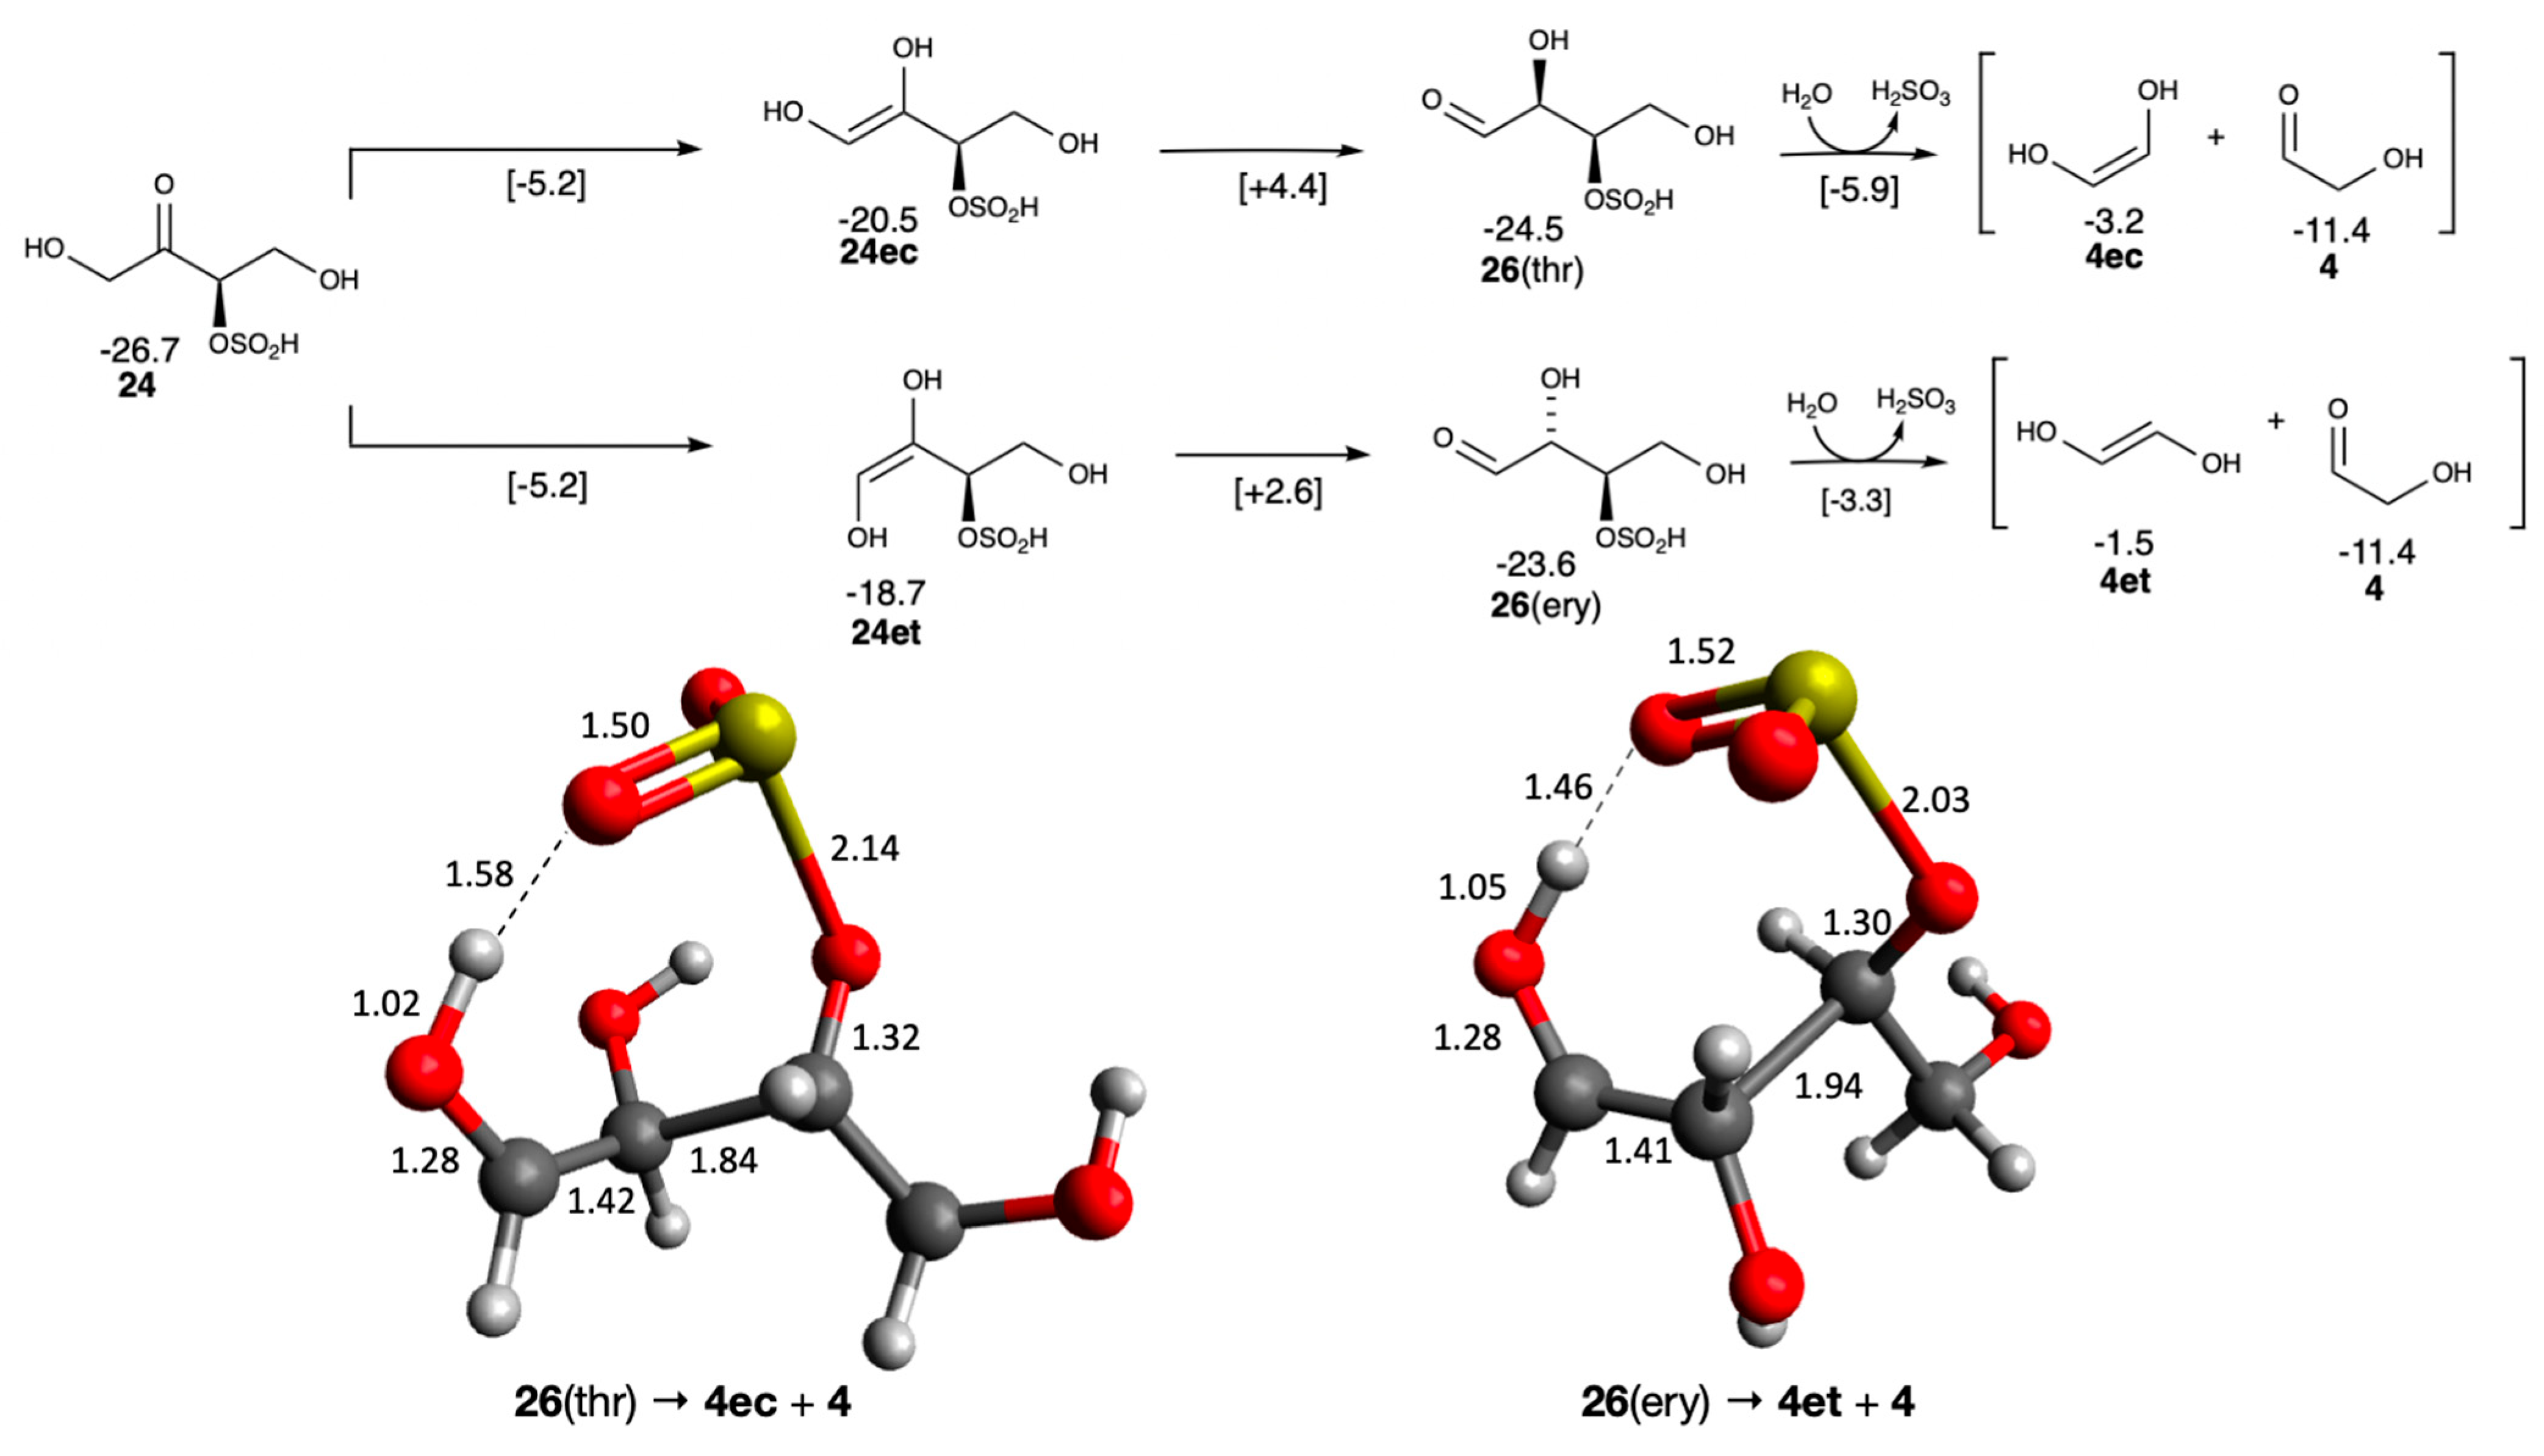
For autocatalysis to occur, two equivalents of glycolaldehyde must be regenerated in the key retro-aldol reaction C4 → 2 C2. The C4 ketose cannot do so without tautomerizing to an aldose, which may be unlikely at higher CH2O concentrations because of competing aldol additions [12]. Also, tautomerization can be slow, and kinetic competition depends on multiple environmental factors. Of the tetroses, only 24 can tautomerize without eliminating the bisulfite group as shown in Figure 10. There are two C4 aldose diastereomers, erythrose and threose, designated 26(ery) and 26(thr) respectively, with threose being marginally more stable by 0.9 kcal/mol. The initial barrier from 24 forming the cis and trans enols is similar in both cases (ΔG‡ = +21.5 kcal/mol), while the second step forming the aldose is marginally more favorable for erythrose compared to threose (the difference in Grel of the two transition states with values of +2.6 and +4.4 kcal/mol). Only the biochemically relevant D-sugars are shown since their L-enantiomers should have similar physical properties.
Figure 10.
Enolization and retro-aldol reaction starting from the 3-bisulfite tetrose (Grel in kcal/mol, bond distances in Å).
Retro-aldol fragmentation of 26 via elimination of the bisulfite moiety leads to glycolaldehyde 4 and its enol (4ec or 4et). Thermodynamically, this is marginally endergonic with ΔG of +2.5 and +3.3 kcal/mol for threose and erythrose, respectively. But the enols eventually convert to 4 (the rate of tautomerization will depend on actual experimental conditions); thus the overall reaction of 26 → 4 + 4 is exergonic with ΔG of −5.7 and −6.6 kcal/mol for threose and erythrose, respectively. In contrast, the non-sulfur analog retro-aldol C4 → 2 C2 is overall endergonic (post-tautomerization) with ΔG of +2.9 and +2.2 kcal/mol for threose and erythrose respectively [33]. In addition, the barrier with intramolecular bisulfite catalysis is modest (ΔG‡ of +18.6 and +20.3 kcal/mol for threose and erythrose respectively) compared to the high barrier ~32 kcal/mol in the non-sulfur analog based on our computational model. The transition states are shown in Figure 10; for threose the breaking C…C bond is 1.84 Å and the breaking S…O bond is 2.14 Å; meanwhile the H from bisulfite has hopped to the aldehyde oxygen. The analogous breaking C…C and S…O bonds are 1.94 Å and 2.03 Å, respectively, in erythrose.
The bisulfite retro-aldol reaction is also more favorable than the thiol analog, the latter being only marginally exergonic (ΔG = −0.2 kcal/mol) also with a high barrier of ~29 kcal/mol [18]. Thus, a key finding of this computational work is that bisulfite adducts lower barriers and make regeneration of the linchpin C2 more favorable under idealized conditions.
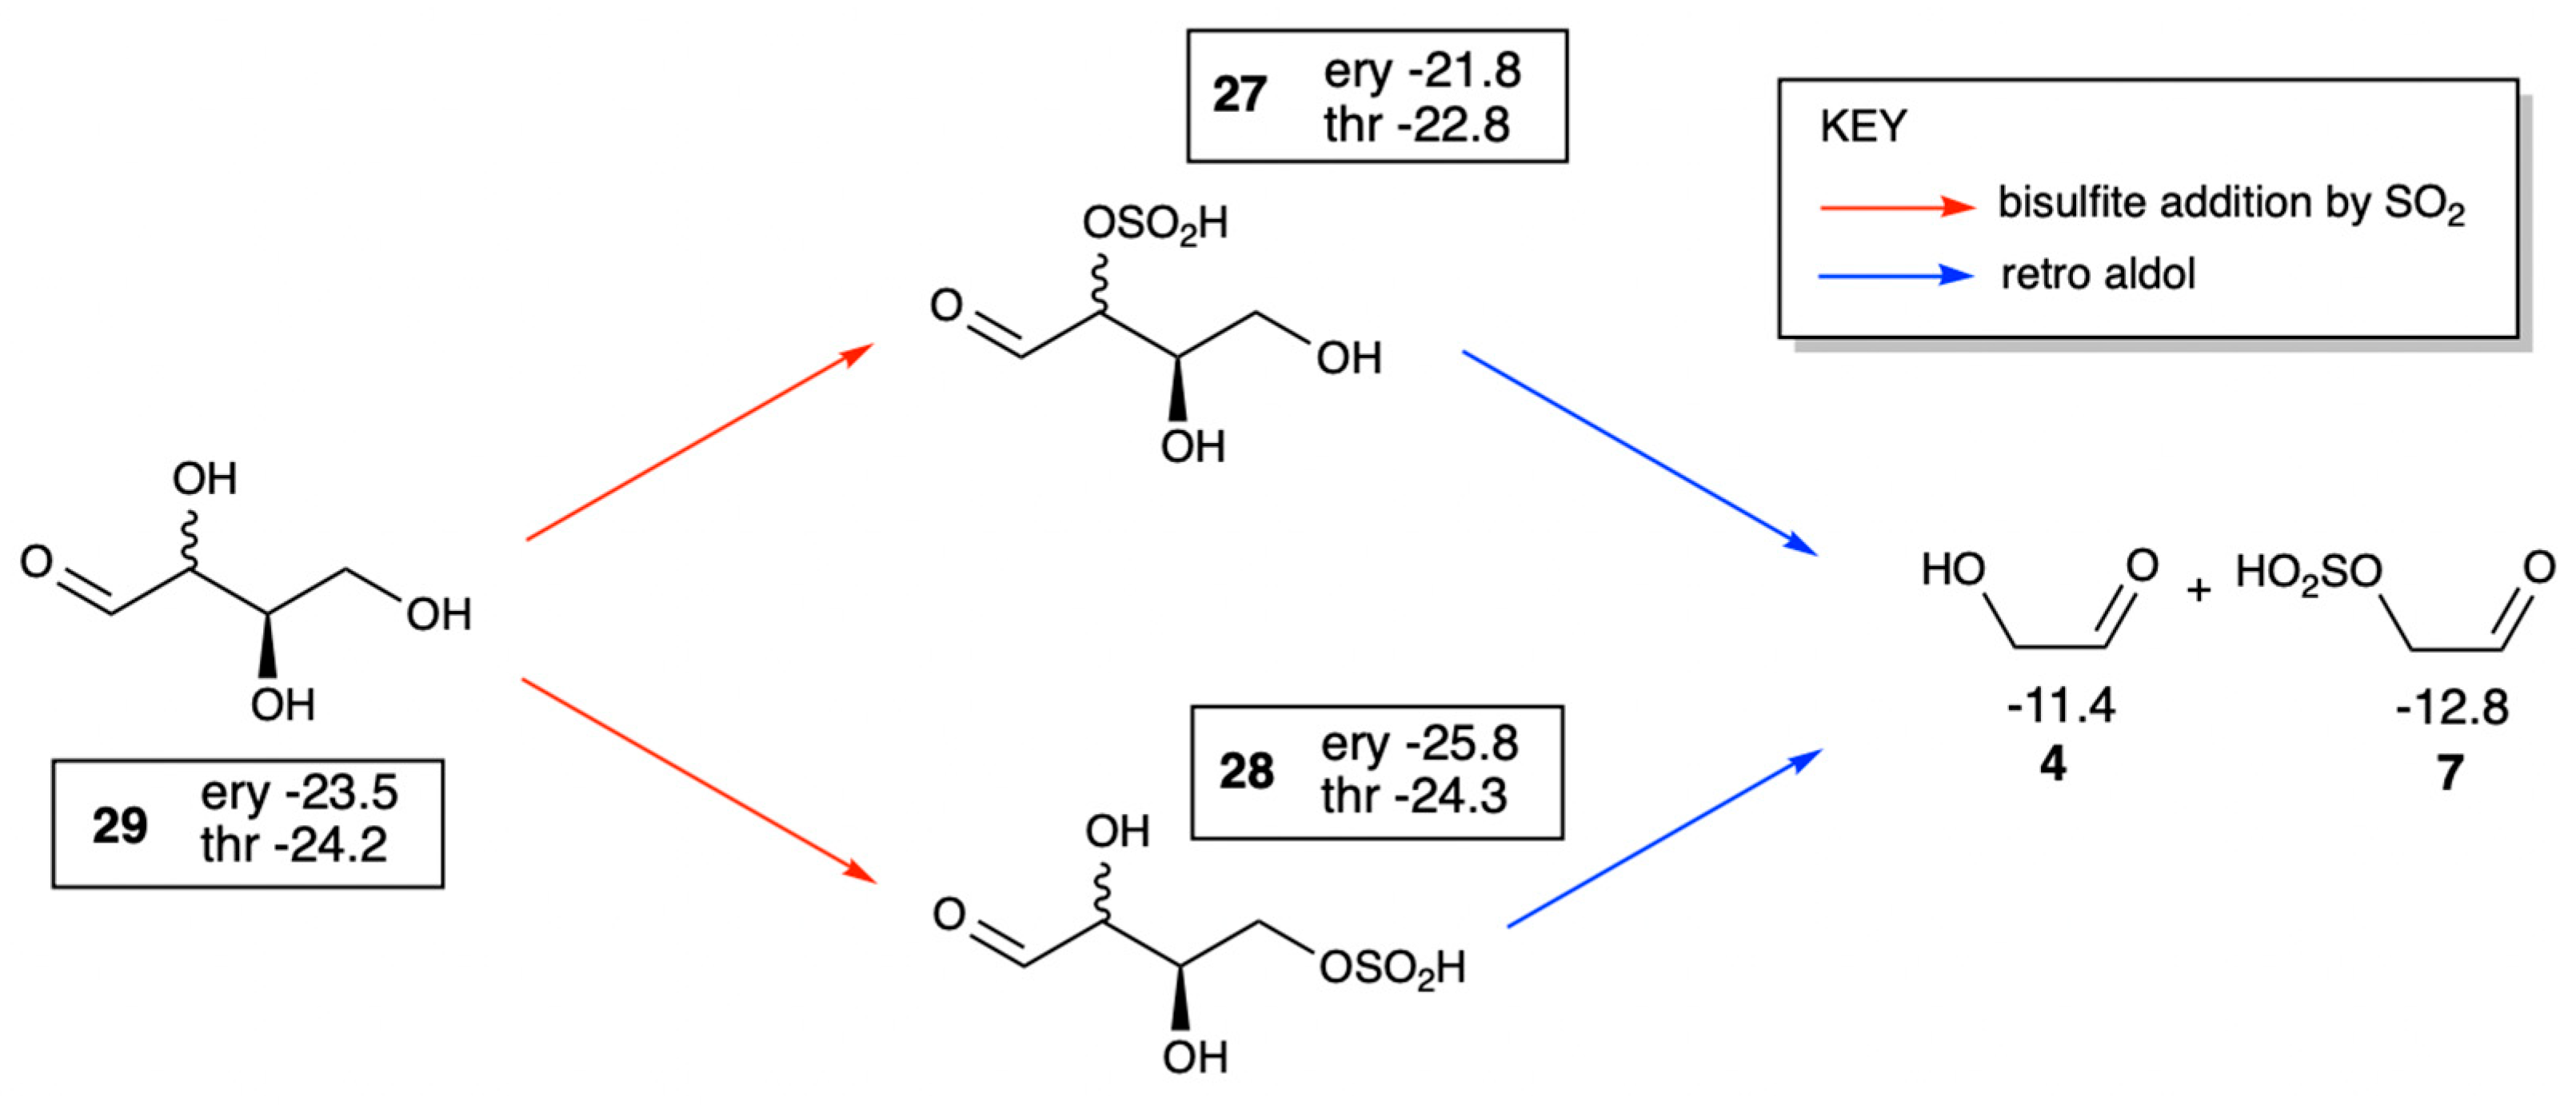
Since the bisulfite moiety can be easily added and removed, we also considered the isomers of adduct 26. As shown in Figure 11, these isomers are 27 and 28 having bisulfite at carbon 2 and carbon 4 respectively. In our model, all three bisulfite adducts (26–28) would be in equilibrium with the non-sulfur analog 29. The threose diastereomer is marginally more stable than erythrose in most cases. Of the three adducts, 28 is the most stable and sterically least hindered. The retro-aldol reactions of 27 and 28 do not require elimination of the bisulfite moiety, and they form the final products 4 and 7. Thermodynamically, the reaction is marginally exergonic for 27 and marginally endergonic for 28 because of the relative stability of 28. Since bisulfite does not act as an intramolecular catalyst in both cases, the barriers are expected to be high similar to the thiol or non-sulfur analogs.
Figure 11.
Enolization and retro-aldol reaction starting from the 3-bisulfite tetrose (Grel in kcal/mol).
The open-chain aldoses can also cyclize to form furanoses. This may sequester the aldose and lengthen its presence in the complex reaction mixture of a formose system. We found no significant effects for the bisulfite adducts, and our results shown in Supplementary Materials are essentially analogous to the non-sulfur analogs [33].
3.6. Thermodynamics of Aldopentose Formation
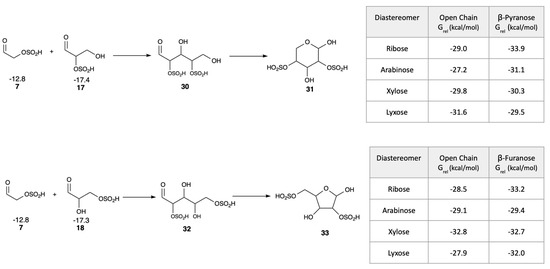
In the Introduction, we discussed experimental work on phosphate adducts, namely that aldol addition of glycolaldehyde phosphate and glyceraldehyde phosphate yielded ribose (with phosphates attached) in high yield over its diastereomers [25]. Could the bisulfite equivalent also favor ribose? Analogous to the experiments, we calculated the free energies of all four D-pentose diastereomers formed in the C2 + C3 → C5 aldol reaction with bisulfite in the 2 and 4 positions. (Mirror image L-enantiomers would have the same energies as their D-counterparts.) As shown in the top half of Figure 12, the reactants 7 and 17 have a combined Grel of −12.8 − 17.4 = −30.2 kcal/mol. Forming the open chain 2,4-pentose adduct 30 is slightly uphill in three cases (and slightly downhill for lyxose with Grel of −31.6 kcal/mol), but ring closure to the pyranose makes the overall reaction thermodynamically favorable for ribose, arabinose and xylose. Thus, our calculations suggest a tendency for cyclization but the actual efficiency will depend on experimental conditions. The 2,4-β-ribopyranose is the most stable diastereomer (Grel of −33.9 kcal/mol), 2.8 kcal/mol more stable than arabinose. Based on our results, ribose appears slightly more stable than other diastereomers, but differences are within computational uncertainty especially comparing arabinose, xylose and lyxose. Comparing the pure sugars (no phosphate or bisulfite) in aqueous solution experimentally, arabinose would be favored over ribose by 0.8 kcal/mol [51]. Thus, the bisulfite moiety may play a similar role in favoring the 2,4-adduct over its diastereomers similar to phosphate.
Figure 12.
Thermodynamics of aldopentose formation and ring closure (Grel in kcal/mol).
For the 2,4-adduct, the bisulfite in the 4-position disfavors furanose formation, making pyranose dominant in our calculations. If instead the bisulfite was in the terminal position of glyceraldehyde (adduct 18), the C2 + C3 aldol addition would yield the 2,5-pentose adduct 32 which can cyclize into the furanose 33 (while being prevented from forming the pyranose because of bisulfite in the 5-position). Forming the 2,5-adduct is slightly exergonic for xylose (Grel of −32.8 kcal/mol) and marginally endergonic in the other three cases although ring closure to the furanose leads to 2,5-β-ribofuranose being thermodynamically the most stable diastereomer (Grel of −33.2 kcal/mol). Xylofuranose, however, is less than 1 kcal/mol higher in energy, thus our results are less clear-cut in this case given the potential computational error.
Overall, our calculations suggest that bisulfite may provide similar thermodynamic stabilization as phosphate, and that ring formation in the aldopentoses helps drive thermodynamic favorability for the aldol addition, but experimental validation is needed. Also, since our calculated Grel values are rather close to each other, this is not an overwhelmingly clear result.
4. Conclusions
Our computational study generating a free energy map for bisulfite adducts of the smallest autocatalytic cycle in the formose system was motivated by the proposal and previous experimental work by Benner and co-workers [31,32]. Within the limitations of our protocol, in our model system, we found that H2SO3 (the hydrate of SO2 in aqueous solution) easily attaches and detaches to both the aldehyde and hydroxyl groups on sugars. The barriers are low and the adducts are in equilibrium with the sugars and have similar Grel values. Since adding the bisulfite moiety to the C1 food molecule (CH2O) is exergonic, this may reduce the effective concentration of free CH2O, thereby decreasing the likelihood of Cannizzaro reactions. This is because the favorable reduction half-reaction is largely compensated by the unfavorable oxidation half-reaction (due to the electron withdrawing bisulfite group on the species being oxidized). Our results also show that the presence of bisulfite increases the barrier by ~5 kcal/mol, kinetically disfavoring the Cannizzaro reaction although actual kinetic effects in complex prebiotic media may vary. This is in good agreement with experimental observations [32]; hence, the presence of SO2 may allow more throughput towards forming sugars and less leakage due to undesired side reactions.
With respect to the aldol additions and key retro-aldol reactions in the simplest formose cycle, our results suggest that intramolecular catalysis by bisulfite lowers the barrier in specific isomers; whether this translates to net autocatalytic flux depends on the relative populations of reactive species. In our optimized transition states, the S…O bond starts to break and the catalytic reaction involves elimination of SO2 from the adduct. A thermodynamic advantage is also observed for the retro-aldol reaction when bisulfite is removed from the tetrose, and the C4 → 2 C2 reaction is now exergonic unlike its non-sulfur analog. Hence, bisulfite adducts might allow for autocatalysis even at lower concentrations of tetrose, unlike the non-sulfur analog where a significant tetrose buildup is needed to overcome the endergonic retro-aldol step. The presence of SO2 might lead to autocatalysis being competitive in the formose system.
The bisulfite adducts of C2 and C3 could subsequently react and stabilize ribose over its diastereomeric counterparts. Computationally, bisulfite may modestly favor ribose over other pentoses, but in aqueous mixtures, this effect may be small relative to competing reactions. Like any other computational study, our results are limited by our protocol. We do not use a high-level density functional and basis set, we apply a simple entropy correction factor, and we use neutral compounds as proxies for what are likely to be anionic species under formose conditions. However, the results from our protocol compare favorably to experimental values in CHO systems, and we hope that extending it to sulfur-analogs will promote further experimental work in CHOS systems as potential proto-metabolites in origins-of-life studies.
Supplementary Materials
The following supporting information can be downloaded at: https://www.mdpi.com/article/10.3390/life16030513/s1, Part 1: Energy breakdown of all molecules in this study, Part 2: Transition state structures, Part 3: Free energy comparison of open tetroses and ring structures, Part 4: XYZ coordinates for transition states.
Author Contributions
E.M.S. and J.K. both contributed to writing (original draft and subsequent edits), formal analysis, investigation, data curation, visualization; J.K. was responsible for conceptualization, methodology, supervision, and project administration. All authors have read and agreed to the published version of the manuscript.
Funding
This research was funded by NASA-Exobiology under Award 80NSSC24K0679.
Institutional Review Board Statement
Not applicable.
Informed Consent Statement
Not applicable.
Data Availability Statement
The data presented in this study are available in the article/Supplementary Materials. Further inquiries can be directed to the corresponding author.
Acknowledgments
This research was supported by the University of San Diego and a research grant from NASA-Exobiology (award # 80NSSC24K0679). Shared computing facilities were provided by the saber3 computing cluster at the University of San Diego.
Conflicts of Interest
The authors declare no conflicts of interest.
References
- Xavier, J.C.; Hordijk, W.; Kauffman, S.; Steel, M.; Martin, W.F. Autocatalytic chemical networks at the origin of metabolism. Proc. Biol. Sci. 2020, 287, 20192377. [Google Scholar] [CrossRef] [Scilit]
- Omran, A.; Menor-Salvan, C.; Springsteen, G.; Pasek, M. The Messy Alkaline Formose Reaction and Its Link to Metabolism. Life 2020, 10, 125. [Google Scholar] [CrossRef] [Scilit]
- Breslow, R. On the Mechanism of the Formose Reaction. Tetrahedron Lett. 1959, 1, 22–26. [Google Scholar] [CrossRef] [Scilit]
- Appayee, C.; Breslow, R. Deuterium studies reveal a new mechanism for the formose reaction involving hydride shifts. J. Am. Chem. Soc. 2014, 136, 3720–3723. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Ricardo, A.; Frye, F.; Carrigan, M.A.; Tipton, J.D.; Powell, D.H.; Benner, S.A. 2-Hydroxymethylboronate as a reagent to detect carbohydrates: Application to the analysis of the formose reaction. J. Org. Chem. 2006, 71, 9503–9505. [Google Scholar] [CrossRef] [Scilit]
- Robinson, W.E.; Daines, E.; van Duppen, P.; de Jong, T.; Huck, W.T.S. Environmental conditions drive self-organization of reaction pathways in a prebiotic reaction network. Nat. Chem. 2022, 14, 623–631. [Google Scholar] [CrossRef] [Scilit]
- van Duppen, P.; Daines, E.; Robinson, W.E.; Huck, W.T.S. Dynamic Environmental Conditions Affect the Composition of a Model Prebiotic Reaction Network. J. Am. Chem. Soc. 2023, 145, 7559–7568. [Google Scholar] [CrossRef] [Scilit]
- Briš, A.; Baltussen, M.G.; Tripodi, G.L.; Huck, W.T.S.; Franceschi, P.; Roithova, J. Direct Analysis of Complex Reaction Mixtures: Formose Reaction. Angew. Chem. Int. Ed. 2024, 63, e202316621. [Google Scholar] [CrossRef] [Scilit]
- Paschek, K.; Kohler, K.; Pearce, B.K.D.; Lange, K.; Henning, T.K.; Trapp, O.; Pudritz, R.E.; Semenov, D.A. Possible Ribose Synthesis in Carbonaceous Planetesimals. Life 2022, 12, 404. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Vinogradoff, V.; Leyva, V.; Mates-Torres, E.; Pepino, R.; Danger, G.; Rimola, A.; Cazals, L.; Serra, C.; Pascal, P.; Meinert, C. Olivine-catalyzed glycolaldehyde and sugar synthesis under aqueous conditions: Application to prebiotic chemistry. Earth Planet. Sci. Lett. 2024, 626, 118558. [Google Scholar] [CrossRef] [Scilit]
- Haas, M.; Lamour, S.; Christ, S.B.; Trapp, O. Mineral-mediated carbohydrate synthesis by mechanical forces in a primordial geochemical setting. Commun. Chem. 2020, 3, 140. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Sutton, S.M.; Pulletikurti, S.; Lin, H.; Krishnamurthy, R.; Liotta, C.L. Abiotic aldol reactions of formaldehyde with ketoses and aldoses—Implications for the prebiotic synthesis of sugars by the formose reaction. Chem 2025, 11, 102553. [Google Scholar] [CrossRef] [Scilit]
- Rappoport, D.; Galvin, C.J.; Zubarev, D.Y.; Aspuru-Guzik, A. Complex Chemical Reaction Networks from Heuristics-Aided Quantum Chemistry. J. Chem. Theory Comput. 2014, 10, 897–907. [Google Scholar] [CrossRef] [Scilit]
- Simm, G.N.; Reiher, M. Context-Driven Exploration of Complex Chemical Reaction Networks. J. Chem. Theory Comput. 2017, 13, 6108–6119. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Roszak, R.; Wołos, A.; Benke, M.; Gleń, Ł.; Konka, J.; Jensen, P.; Burgchardt, P.; Zadlo-Dobrowolska, A.; Janiuk, P.; Szymkuc, S.; et al. Emergence of metabolic-like cycles in blockchain-orchestrated reaction networks. Chem 2024, 10, 952–970. [Google Scholar] [CrossRef] [Scilit]
- Kan, H.W.; Li, X.T.; Zhang, J.Z.H. Mechanistic insights into formose side reactions: Cannizzaro disproportionation and competing pathways. J. Chem. Phys. 2025, 163, 085102. [Google Scholar] [CrossRef] [Scilit]
- Kua, J.; Avila, J.E.; Lee, C.G.; Smith, W.D. Mapping the Kinetic and Thermodynamic Landscape of Formaldehyde Oligomerization under Neutral Conditions. J. Phys. Chem. A 2013, 117, 12658–12667. [Google Scholar] [CrossRef] [Scilit]
- Kua, J.; Peña, M.T.; Cotter, S.N.; Leca, J. Sulfur Analogs of the Core Formose Cycle: A Free Energy Map. Life 2025, 15, 1. [Google Scholar] [CrossRef] [Scilit]
- Neveu, M.; Kim, H.J.; Benner, S.A. The “strong” RNA world hypothesis: Fifty years old. Astrobiology 2013, 13, 391–403. [Google Scholar] [CrossRef] [Scilit]
- Ricardo, A.; Carrigan, M.A.; Olcott, A.N.; Benner, S.A. Borate minerals stabilize ribose. Science 2004, 303, 196. [Google Scholar] [CrossRef] [Scilit]
- Springsteen, G.; Joyce, G.F. Selective derivatization and sequestration of ribose from a prebiotic mix. J. Am. Chem. Soc. 2004, 126, 9578–9583. [Google Scholar] [CrossRef] [Scilit]
- Lambert, J.B.; Gurusamy-Thangavelu, S.A.; Ma, K. The silicate-mediated formose reaction: Bottom-up synthesis of sugar silicates. Science 2010, 327, 984–986. [Google Scholar] [CrossRef] [Scilit]
- Usami, K.; Okamoto, A. Hydroxyapatite: Catalyst for a one-pot pentose formation. Org. Biomol. Chem. 2017, 15, 8888–8893. [Google Scholar] [CrossRef] [Scilit]
- Zubay, G. Studies on the lead-catalyzed synthesis of aldopentoses. Orig. Life Evol. Biosph. 1998, 28, 13–26. [Google Scholar] [CrossRef] [Scilit]
- Mueller, D.; Pitsch, S.; Kittaka, A.; Wagner, E.; Wintner, C.E.; Eschenmoser, A. Chemistry of a-aminonitriles. Aldomerization of glycolaldehyde phosphate to rac-hexose 2,4,6-triphosphates and (in presence of formaldehyde) rac-pentose 2,4-diphosphates: Rac-allose 2,4,6-triphosphate and rac-ribose 2,4-diphosphate are the main reaction products. Helv. Chim. Acta 1990, 73, 1410–1468. [Google Scholar] [CrossRef] [Scilit]
- Camprubi, E.; Harrison, S.A.; Jordan, S.F.; Bonnel, J.; Pinna, S.; Lane, N. Do Soluble Phosphates Direct the Formose Reaction towards Pentose Sugars? Astrobiology 2022, 22, 981–991. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Gulick, A. Phosphorus as a factor in the origin of life. Am. Sci. 1955, 43, 479–489. [Google Scholar]
- Keefe, A.D.; Miller, S.L. Are polyphosphates or phosphate esters prebiotic reagents? J. Mol. Evol. 1995, 41, 693–702. [Google Scholar] [CrossRef] [Scilit]
- Brady, M.P.; Tostevin, R.; Tosca, N.J. Marine phosphate availability and the chemical origins of life on Earth. Nat. Commun. 2022, 13, 5162. [Google Scholar] [CrossRef] [Scilit]
- Trail, D.; Watson, E.B.; Tailby, N.D. The oxidation state of Hadean magmas and implications for early Earth’s atmosphere. Nature 2011, 480, 79–82. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Benner, S.A.; Kim, H.J.; Biondi, E. Prebiotic Chemistry that Could Not Not Have Happened. Life 2019, 9, 84. [Google Scholar] [CrossRef] [Scilit]
- Kawai, J.; McLendon, D.C.; Kim, H.J.; Benner, S.A. Hydroxymethanesulfonate from Volcanic Sulfur Dioxide: A “Mineral” Reservoir for Formaldehyde and Other Simple Carbohydrates in Prebiotic Chemistry. Astrobiology 2019, 19, 506–516. [Google Scholar] [CrossRef] [Scilit]
- Kua, J.; Tripoli, L.P. Exploring the Core Formose Cycle: Catalysis and Competition. Life 2024, 14, 933. [Google Scholar] [CrossRef] [Scilit]
- Kua, J.; Hanley, S.W.; De Haan, D.O. Thermodynamics and Kinetics of Glyoxal Dimer Formation: A Computational Study. J. Phys. Chem. A 2008, 112, 66–72. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Kua, J.; Galloway, M.M.; Millage, K.D.; Avila, J.E.; De Haan, D.O. Glycolaldehyde Monomer and Oligomer Equilibria in Aqueous Solution: Comparing Computational Chemistry and NMR Data. J. Phys. Chem. A 2013, 117, 2997–3008. [Google Scholar] [CrossRef] [Scilit]
- Kua, J.; Hernandez, A.L.; Velasquez, D.N. Thermodynamics of Potential CHO Metabolites in a Reducing Environment. Life 2021, 11, 1025. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Vosko, S.H.; Wilk, L.; Nusair, M. Accurate spin-dependent electron liquid correlation energies for local spin density calculations: A critical analysis. Can. J. Phys. 1980, 58, 1200–1211. [Google Scholar] [CrossRef] [Scilit]
- Becke, A.D. Density-functional exchange-energy approximation with correct asymptotic behavior. Phys. Rev. A 1988, 38, 3098–3100. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Lee, C.; Yang, W.; Parr, R.G. Development of the Colle-Salvetti correlation-energy formula into a functional of the electron density. Phys. Rev. B 1988, 37, 785–789. [Google Scholar] [CrossRef] [Scilit]
- Becke, A.D. Density-functional thermochemistry. III. The role of exact exchange. J. Chem. Phys. 1993, 98, 5648–5652. [Google Scholar] [CrossRef] [Scilit]
- Marenich, A.V.; Cramer, C.J.; Truhlar, D.G. Universal solvation model based on solute electron density and on a continuum model of the solvent defined by the bulk dielectric constant and atomic surface tensions. J. Phys. Chem. B 2009, 113, 6378–6396. [Google Scholar] [CrossRef] [Scilit]
- Warshel, A.; Florian, J. Computer simulations of enzyme catalysis: Finding out what has been optimized by evolution. Proc. Natl. Acad. Sci. USA 1998, 95, 5950–5955. [Google Scholar] [CrossRef] [Scilit]
- Wiberg, K.B.; Bailey, W.F. Chiral diamines 4: A computational study of the enantioselective deprotonation of Boc-pyrrolidine with an alkyllithium in the presence of a chiral diamine. J. Am. Chem. Soc. 2001, 123, 8231–8238. [Google Scholar] [CrossRef] [Scilit]
- Nielsen, R.J.; Keith, J.M.; Stoltz, B.M.; Goddard, W.A., 3rd. A computational model relating structure and reactivity in enantioselective oxidations of secondary alcohols by (−)-sparteine-Pd(II) complexes. J. Am. Chem. Soc. 2004, 126, 7967–7974. [Google Scholar] [CrossRef] [Scilit]
- Deubel, D.V.; Lau, J.K. In silico evolution of substrate selectivity: Comparison of organometallic ruthenium complexes with the anticancer drug cisplatin. Chem. Comm. 2006, 23, 2451–2453. [Google Scholar] [CrossRef] [Scilit]
- Wertz, D.H. Relationship between the gas-phase entropies of molecules and their entropies of solvation in water and 1-octanol. J. Am. Chem. Soc. 1980, 102, 5316–5322. [Google Scholar] [CrossRef] [Scilit]
- Abraham, M.H. Relationship between solution entropies and gas phase entropies of nonelectrolytes. J. Am. Chem. Soc. 1981, 103, 6742–6744. [Google Scholar] [CrossRef] [Scilit]
- Krizner, H.E.; De Haan, D.O.; Kua, J. Thermodynamics and Kinetics of Methylglyoxal Dimer Formation: A Computational Study. J. Phys. Chem. A 2009, 113, 6994–7001. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Damer, B.; Deamer, D. The Hot Spring Hypothesis for an Origin of Life. Astrobiology 2020, 20, 429–452. [Google Scholar] [CrossRef] [Scilit]
- Kim, H.J.; Ricardo, A.; Illangkoon, H.I.; Kim, M.J.; Carrigan, M.A.; Frye, F.; Benner, S.A. Synthesis of Carbohydrates in Mineral-Guided Prebiotic Cycles. J. Am. Chem. Soc. 2011, 133, 9457–9468. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Tewari, Y.B.; Goldberg, R.N. An investigation of the equilibria between aqueous ribose, ribulose, and arabinose. Biophys. Chem. 1985, 22, 197–204. [Google Scholar] [CrossRef] [Scilit] [PubMed]
Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2026 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license.