


Abstract
Generic polymer models capturing the chain connectivity and the non-bonded excluded-volume interactions between polymer segments can be classified into hard- and soft-core models depending on their non-bonded pair potential. Here we compared the correlation effects on the structural and thermodynamic properties of the hard- and soft-core models given by the polymer reference interaction site model (PRISM) theory, and found different behaviors of the soft-core models at large invariant degree of polymerization (IDP) depending on how IDP is varied. We also proposed an efficient numerical approach, which enables us to accurately solve the PRISM theory for chain lengths as large as 106.
While they do not correspond to any chemically specific polymer, generic polymer models are widely used in theoretical and simulation studies in the field of polymer physics as they capture two essential features of all polymers: chain connectivity and non-bonded excluded-volume interactions. Compared to atomistic models that can represent specific polymers used in experiments, molecular simulations of generic models can reach much larger length scales and much longer time scales, and theoretical studies of generic models can also be performed. Depending on whether or not the excluded-volume interactions in generic models prevent complete overlapping of polymer segments, they can be classified into hard-core models (such as those based on the hard-sphere chain model, the Kremer–Grest model [1], and the various self- and mutual-avoiding walk models on a lattice) and soft-core models (such as those used in the dissipative particle dynamics (DPD) simulation [2], fast Monte Carlo simulations [3,4,5,6,7], field-theoretic simulation (FTS) [8], variants of FTS under the partial saddle-point approximation [9], single-chain-in-mean-field simulation [10] and hybrid particle field molecular dynamics simulation [11] both under the quasi-instantaneous field approximation [10]). Taking the study of polymer melts as an example, while hard-core models have been used in conventional molecular simulations for a long time, they have the disadvantage that their chain lengths N used in such simulations (as limited by the computational cost) are too short compared to those in typical experiments; in other words, such conventional simulations significantly exaggerate the fluctuations in polymer melts compared to experiments [6,7,12]. In contrast, simulations of the more recently proposed soft-core models can readily reach the extent of fluctuations in typical experiments by increasing the chain number density (or equivalently the segment number density ρ at finite N) instead of N [6,7,12].
In this Letter we focus on a simple but important class of generic models for compressible homopolymer melts (or equivalently homopolymer solutions in an implicit solvent) in the continuum, with the excluded-volume interaction between polymer segments described by a short-range, isotropic and purely repulsive pair potential βunb(r), where β ≡ 1/kBT with kB being the Boltzmann constant and T the thermodynamic temperature of the system; this is the basis of more complicated polymer models having attractions and/or more species. The hard- and soft-core models can then be classified according to whether diverges or not. This classification becomes clear after we write the total dimensionless non-bonded interaction energy for a system of n chains each having N segments in volume V under the commonly used pairwise additivity as
where ri denotes the spatial position of the ith segment in the system, is the segment volume fraction at r, and the last term deducting the self-interaction of segments gives an unimportant constant; while molecular simulations of this system can be performed at finite ρ ≡ nN/V for any βunb(r) (along with a chain-connectivity model), for a homogeneous system (i.e., ) the widely used polymer self-consistent field (SCF) theory [13] gives the dimensionless internal energy per chain due to the non-bonded interaction (due to its mean-field approximation that neglects the system fluctuations and correlations), which diverges if does (i.e., for the hard-core models). It is then clear that the SCF theory can only be applied to soft-core models, where one can define the dimensionless excluded-volume interaction parameter ε > 0 via unb(r) = εu0(r) with the normalized pair potential u0(r) satisfying . Another necessary condition for applying the SCF theory (i.e., having finite βUnb/n) is ε ∝ ρ−1.
Here we compare the correlation effects on the structural and thermodynamic properties of hard- and soft-core generic polymer models, which has rarely been reported [14], to further reveal their differences. For this purpose we choose the polymer reference interaction site model (PRISM) theory proposed by Schweizer and Curro [15], which has been applied to many polymeric systems, including homopolymer melts, solutions, blends, block copolymers, nanocomposites, polyelectrolytes, etc. [16,17,18,19] It can be considered as the most successful molecular-level theory to date for studying the correlations in homogeneous polymeric systems.
For the above homopolymer melts, the PRISM equation is given by
where is the interchain total segment pair correlation function (PCF) with and σ the segment diameter (i.e., the range of βunb(r)), is the normalized (i.e., ) intrachain segment PCF, is the interchain direct segment PCF, denotes the 3D Fourier transform of a radial function with q being the wavenumber (in units of 1/σ), and is the dimensionless segment number density. For given N, and ω, to solve for both h and c, a closure providing an approximate relation between them is needed; here we take the commonly used Percus–Yevick (PY) closure [20]
which works well for our class of generic models where is short-range and purely repulsive.
To be more specific, we consider two commonly used generic polymer models: the tangent hard-sphere chain (THSC) and the DPD models; the former is a hard-core model that uses with specifying the dimensionless bonded potential between two adjacent segments on the same chain and the hard-sphere (HS) potential for and 0 otherwise as , and the latter is a soft-core model that uses and the DPD potential for and 0 otherwise as with the dimensionless interaction parameter chosen to mimic the compressibility of water [2]. In the thermodynamic limit, the structural and thermodynamic properties of these two models are controlled only by N and ; typically, molecular simulations of the DPD model uses or 5.
Finally, we note that is needed as an input for PRISM calculations. While in general the chain conformations characterized by depend on , for simplicity here we use the ideal-chain conformations by setting to with for the THSC model and for the DPD model, where , q is the wave vector for the 3D Fourier transform, and q=|q|.
For the two generic models that we consider here, the PY closure gives
. Since all previously reported numerical methods for PRISM calculations [21,22,23,24] are not optimal in this case, we first propose an efficient numerical approach as follows. We uniformly discretize the real-space interval [0, 1] into m subintervals (thus
into
subintervals) each of length
, where
denotes the real-space cut-off, and take
(i = 0, …, m−1 for the DPD model and i = 0, …, m with
for the THSC model) as the independent variables to be solved. Our approach has three steps:
- Given the initial guess of the independent variables and , for the DPD model we calculate (j = 1, …, M−1) via the discrete sine transform of type I (DST) [25], which has the computational complexity of O(MlnM) and gives , the reciprocal-space cut-off qc = qM = mπ and ; for the THSC model, due to the discontinuities in both and its 1st-order derivative at , we use an auxiliary function for and otherwise with and (calculated via the fourth-order backward finite-difference formula [26]), which is continuous in both its value and 1st-order derivative, to calculate (j = 1, …, M) via the DST, which gives . We also calculate for the DPD model and for the THSC model via the Romberg integration (RI) [27].
- We calculate (j = 0, …, M) obtained from Equation (1) with being the interchain indirect segment PCF (note that for the DPD model while for the THSC model), then for the DPD model (j = 1, …, M−1) via the DST (which gives ); for the THSC model, we use another auxiliary function to calculate (j = 1, …, M) via the DST, which gives . We also calculate for both models via the RI.
- We calculate (i = 0, …, m–1 for the DPD model and i = 0, …, m for the THSC model), then use the residual errors of Equation (2) (which becomes for the THSC model) to converge the independent variables via the Anderson mixing [28], which has the computational complexity of O(m) and can quickly converge a large set of nonlinear equations to a high accuracy.
We use the convergence criterion of εc < 10−10 with εc denoting the maximum absolute value of the residual errors of the PY closure over all (i = 0, …, m–1 for the DPD model and i = 0, …, m for the THSC model), and choose the values of m (=4096 for the THSC model and 512 for the DPD model) and ( if N < 100 and otherwise, rounded to the nearest integer, to capture the correlation-hole effect [29]) such that the discretization errors are comparable to εc. Our numerical approach has the least number of independent variables to be iteratively solved, greatly reduces m (thus M) both by analytically treating the discontinuities in the THSC model and by taking the inverse Fourier transform only for (which decays toward 0 with increasing q much faster than both and ), and is essential for us to accurately solve the PRISM-PY theory for N as large as 106 (where for the THSC model M is about 8.2 × 10⁶!). To the best of our knowledge, analytically treating the discontinuities caused by the HS potential has not been reported in numerical calculations of even the widely studied Ornstein–Zernike (OZ) equation [30] (to which Equation (1) reduces for N = 1); in Supplemental Material we show that our numerical approach gives several orders of magnitude more accurate results than pyPRISM [19], a recently developed Python-based open-source framework for PRISM calculations.
In the limit of N→∞ and σ→0 at finite root-mean-square end-to-end distance of the ideal chain , the THSC model becomes the hard-core Gaussian thread model [31] (HC CGC-δ, where Re,0 is taken as the unit of length); to compare the PRISM-PY results of these two models, we define two dimensionless parameters: and the invariant degree of polymerization [32]
;
controls the fluctuations in polymer melts, and for the THSC model it is easy to show that
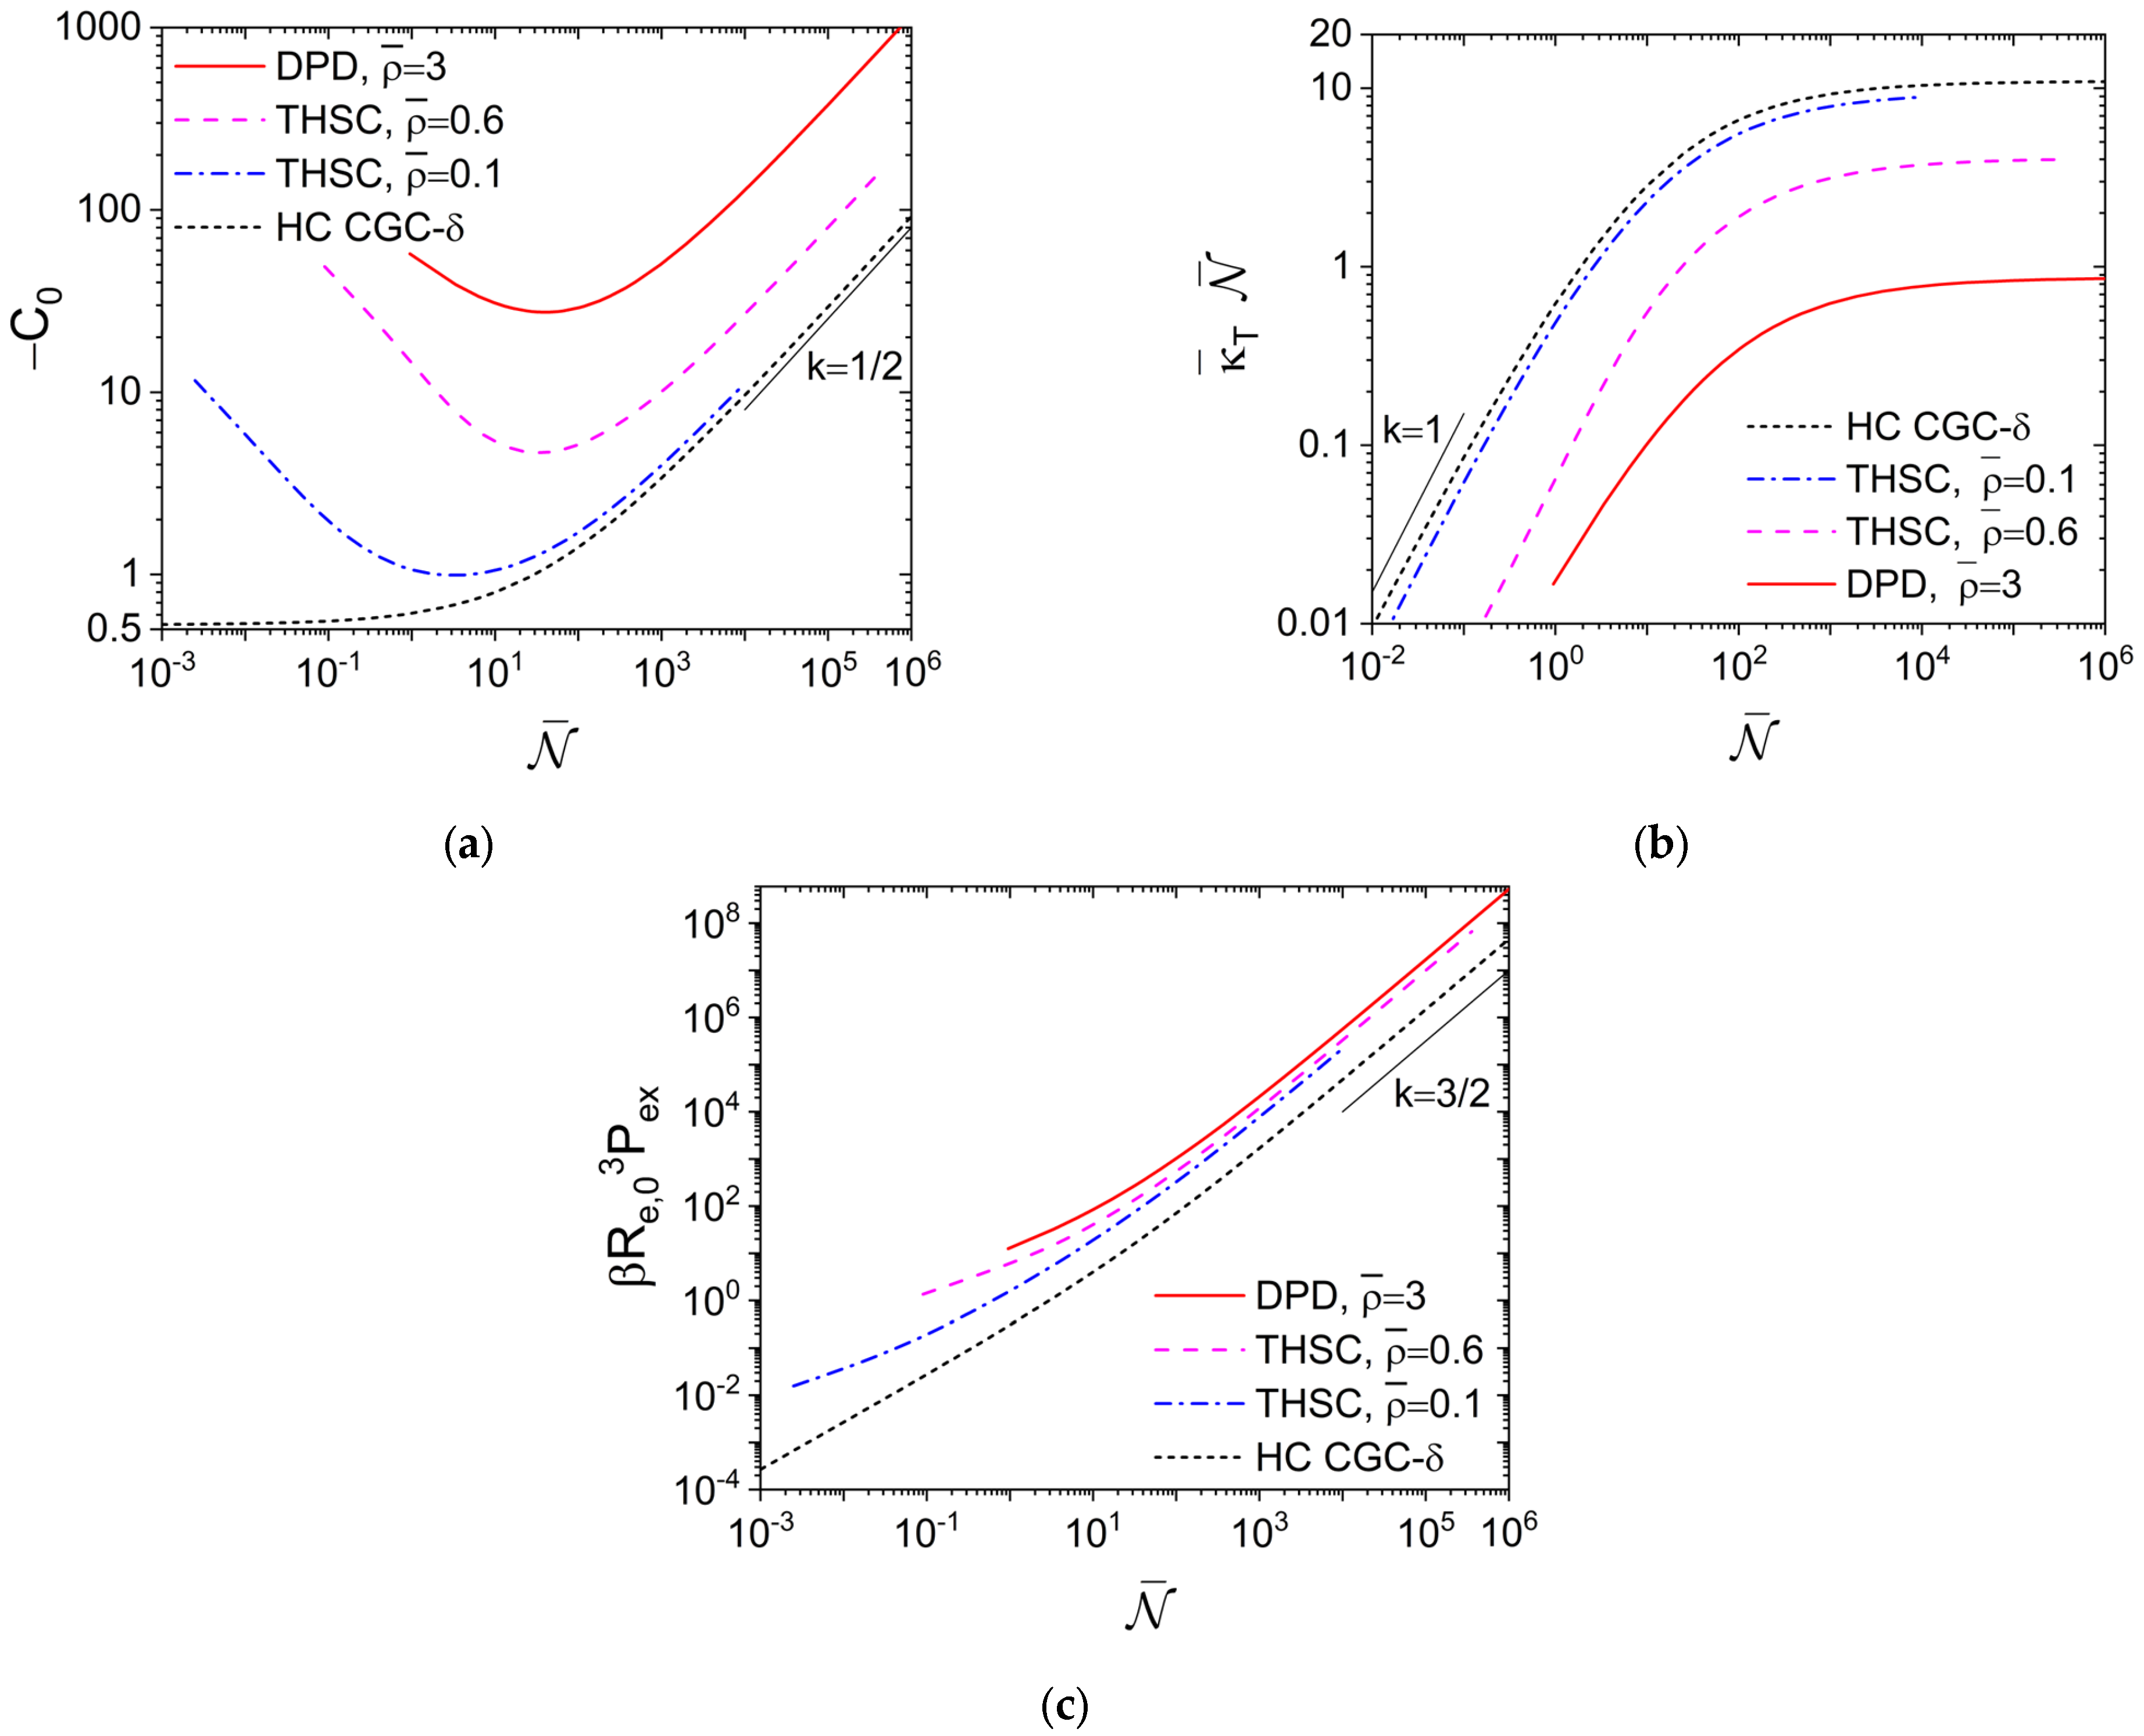
∝ N at large N. Figure 1a shows how C0 varies with
for the THSC and HC CGC-δ models; for the latter model,
is the only parameter, the PRISM-PY equation is given by Equation (18) in our previous work [14] and the corresponding numerical results for
≥ 100 are shown in figure 8b there. We see that, while −C0 increases monotonically with increasing
for the HC CGC-δ model, it exhibits a minimum for the THSC model. At given
due to its N→∞ the HC CGC-δ model corresponds to the limit of
of the THSC model as implied in Figure 1a. At large
, we see that
in all cases. This is in accordance with an asymptotic value of at given for the THSC model, while for the HC CGC-δ model. Figure 1a also shows that the DPD model at gives qualitatively the same behavior of C0 vs.
as that for the THSC model.
Figure 1.
Logarithmic plot of (a) , (b) normalized isothermal compressibility , and (c) excess (virial) pressure due to the interchain repulsion Pex vs. the invariant degree of polymerization
of various models. The k-value gives the slope of the corresponding straight line. See the main text for more details.
With the normalized isothermal compressibility
given by the compressibility equation, where ρc ≡ n/V is the chain number density and is the isothermal compressibility with P denoting the system pressure, Figure 1b presents essentially the same data as in Figure 1a, but in a way that can be compared with real polymers used in experiments. As shown in figure 2 of our previous work [14],
for polyethylene (at 180 °C) and 0.119 for polystyrene (at 280 °C), independent of their
≥103; this is consistent with
at large
shown in Figure 1a. On the other hand, while
is expected for very small
, the smallest
(given by N = 2) is about 0.0025, 0.090 and 0.95, respectively, for the THSC model at and 0.6 and the DPD model at . Clearly, both hard- and soft-core models can be used to describe the excluded-volume interactions in real polymers, and experimental values of can be achieved by adjusting , for example, in the THSC and DPD models. We attribute the largest at the same
given by the HC CGC-δ model to its σ→0, and note that the DPD model at is actually “harder” (i.e., more difficult to compress) than the hard-core models studied here.
Figure 1c shows that at large
, the dimensionless excess (virial) pressure due to the interchain repulsion
scales with
for the THSC model; this is due to the same scaling of Re,03 with N and also found for the HC CGC-δ model (where
). We also see that the HC CGC-δ model gives a much smaller βRe,03Pex than the THSC model at the same
, again due to its σ→0. Figure 1c further shows that at large
, the DPD model at gives the same scaling of
with
as the hard-core models; at the same
, it has even the largest , in accordance with its smallest shown in Figure 1b.
Note that for both the THSC and DPD models,
is varied by changing N at fixed in Figure 1, which makes
and N to be approximately on the same order; it is therefore very difficult, if possible at all, to reach via this way in molecular simulations even a relatively small
-value of 104 used in experiments. As aforementioned, molecular simulations of soft-core models can readily reach
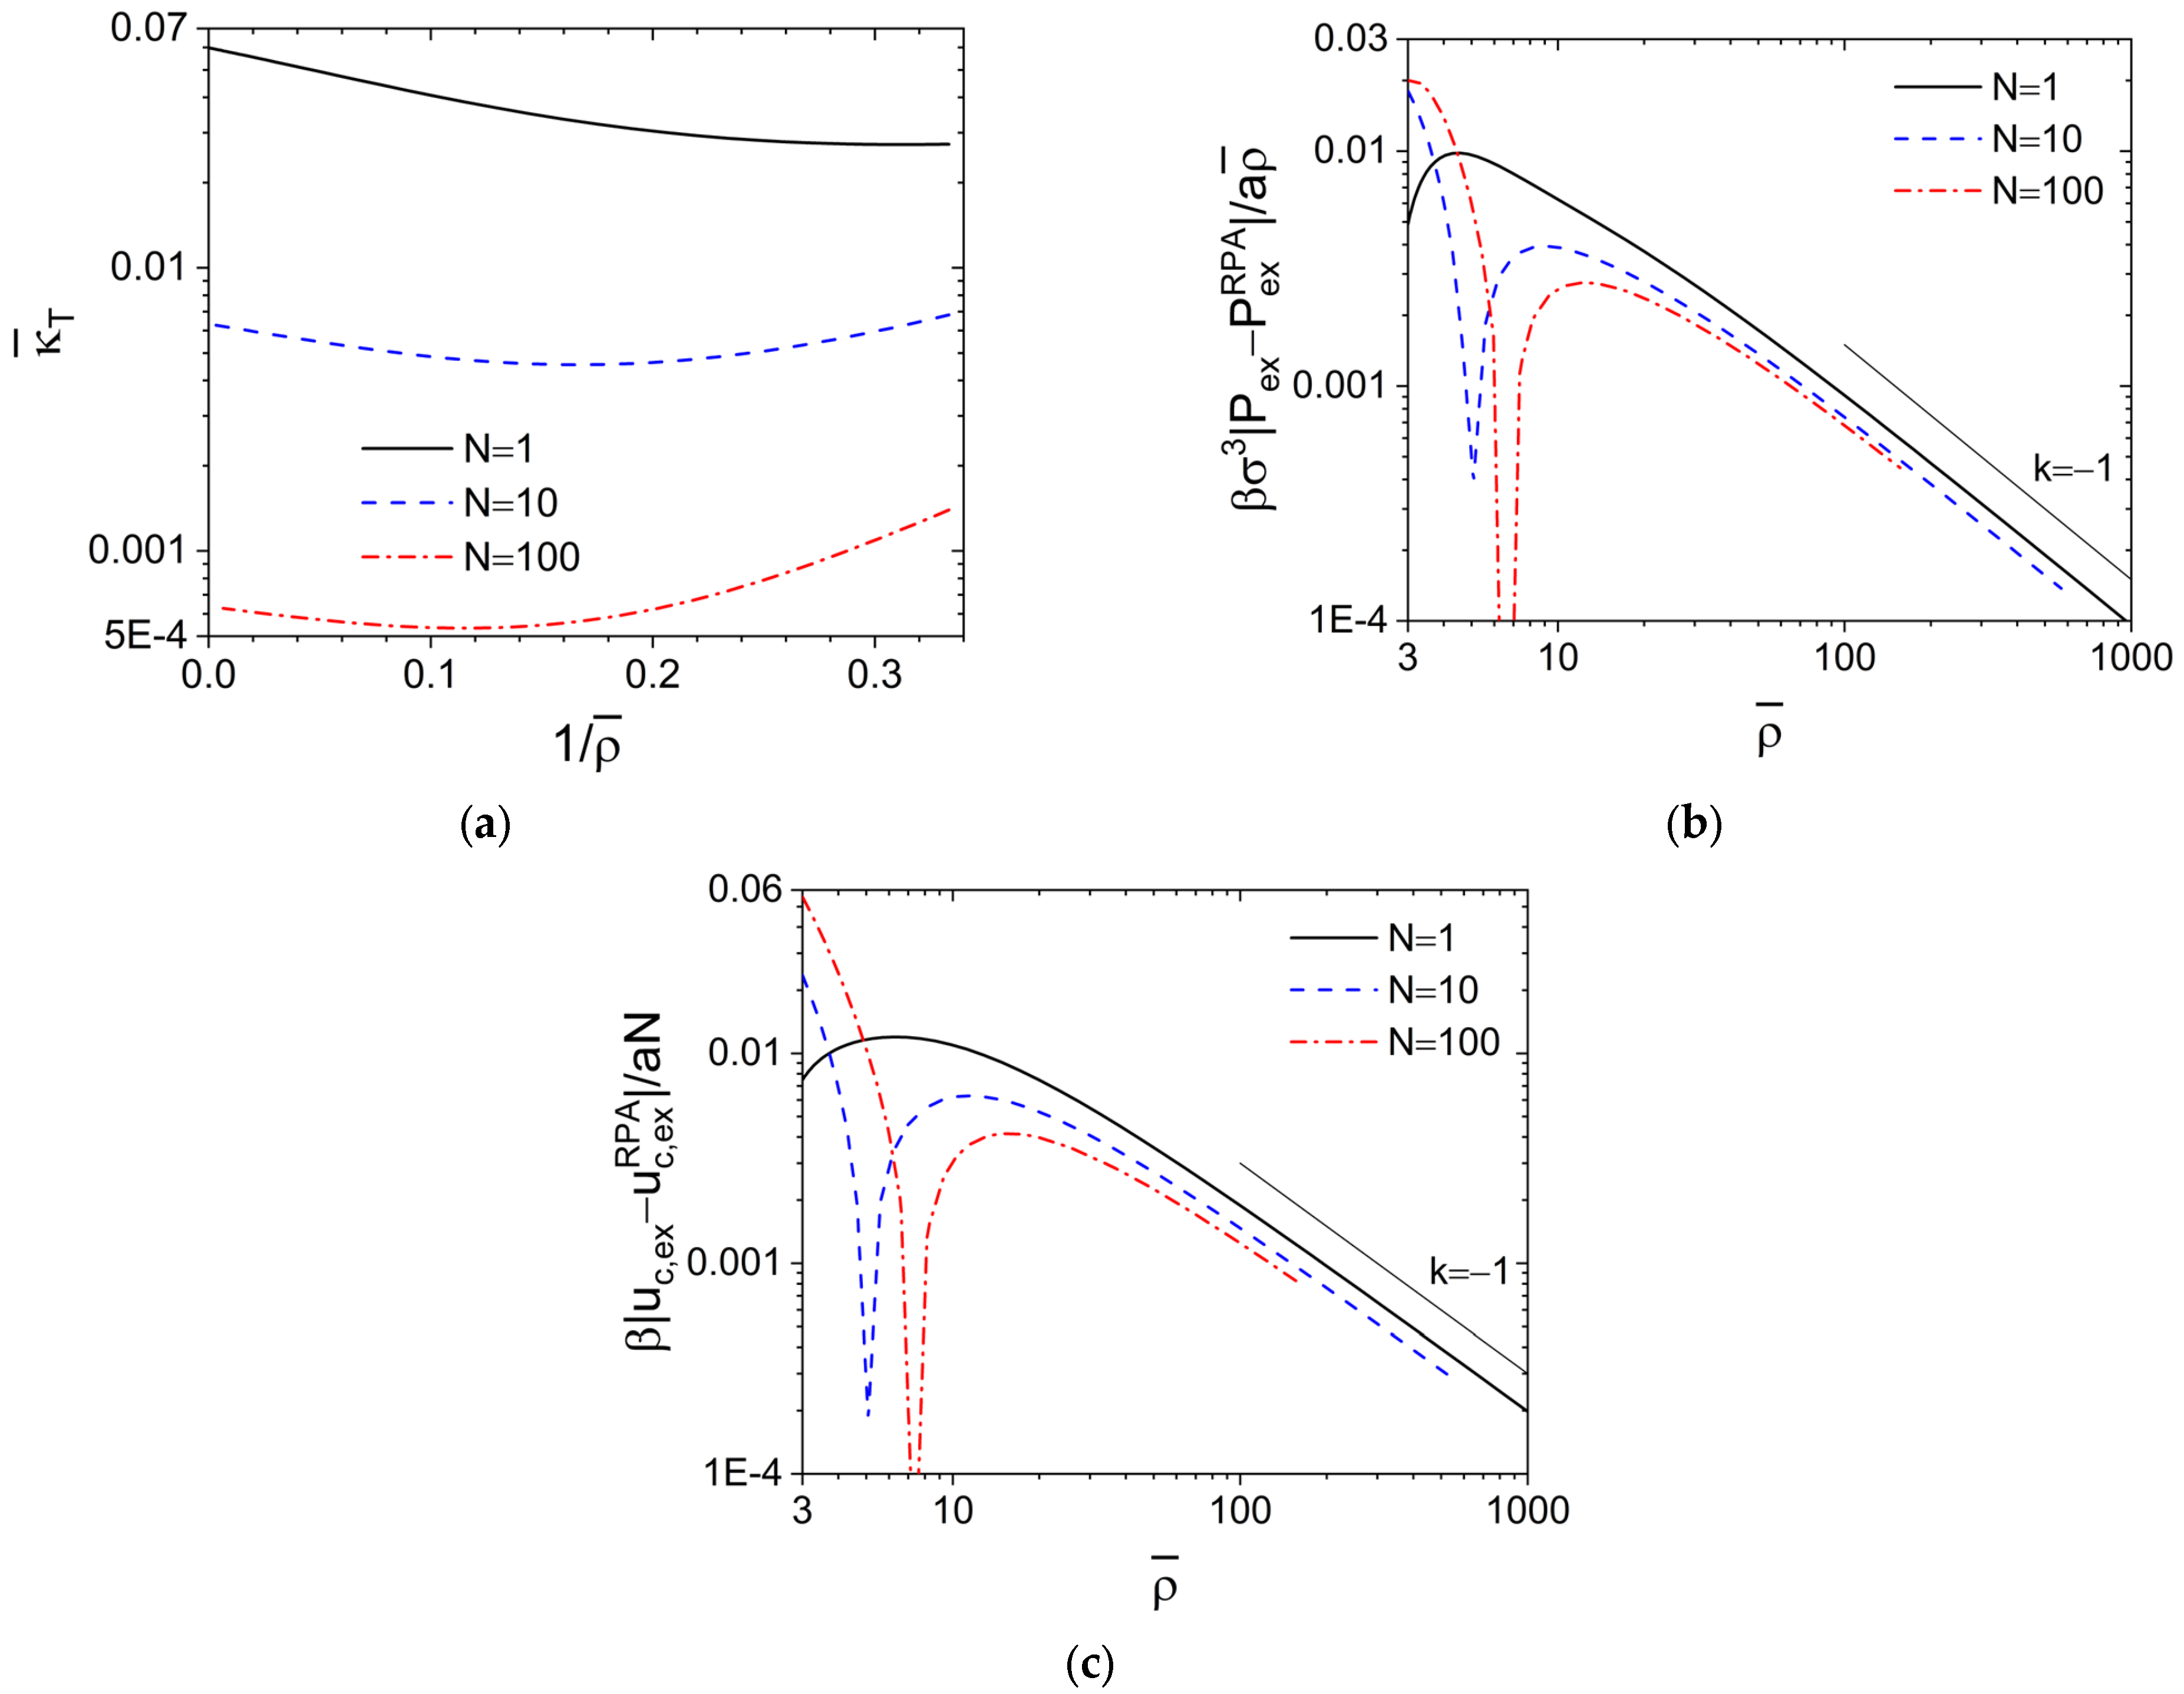
-values used in experiments by increasing at fixed N [6,7,12]. For the DPD model at large , and the PY closure approaches the random-phase approximation (RPA) closure [33,34] , which gives and independent of N. We then obtain from the compressibility equation. Figure 2a shows vs. obtained via the compressibility equation from our PRISM-PY calculations of the DPD model at various N, where each curve exhibits a minimum with its location shifting to smaller with increasing N and the intercept of each (extrapolated) curve with the left axis (i.e., in the limit of ) gives the corresponding . Clearly, the difference between and is entirely due to that between the PY and RPA closures.
Figure 2.
(a) Semi-logarithmic plot of the normalized isothermal compressibility
and logarithmic plot of the difference between (b) the excess (virial) pressure Pex and (c) the excess internal energy per chain uc,ex due to the interchain interactions given by the PRISM-PY calculations and that by the PRISM-RPA calculations of the DPD model at various chain lengths N. The k-value gives the slope of the corresponding straight line. See the main text for more details.
A C0 vs.
plot (not shown) can be obtained from Figure 2a for the DPD model. In particular, the RPA closure gives
, indicating that
at large
; this is in clear contrast to
for the hard-core models and the DPD model at shown in Figure 1a, but consistent with the soft-core Gaussian thread (SC CGC-δ) model (which is equivalent to the well-known Edwards model [35]) shown in figure 8a of our previous work [14], where N→∞ and σ→0 at finite Re,0 and is used with a finite dimensionless parameter controlling the repulsion strength between polymer segments. The behavior of soft-core models at large
, therefore, depends on how
is varied, i.e., whether by changing N at fixed (thus the excluded-volume interaction parameter ε is fixed) or by changing at fixed N (thus ε is also varied as ); in the former case correlations exist even in the limit of N→∞, while in the latter case the SCF theory becomes exact in the limit of (at finite N) where neither fluctuations nor correlations exist.
As aforementioned, with increasing at fixed N, the PY closure approaches the RPA closure, which gives thus according to Equation (1). In the limit of , we have and , thus the SCF results of and independent of N, where denotes the dimensionless excess internal energy per chain due to the interchain repulsion. On the other hand, the differences between the SCF and RPA results as given by and are independent of .
Finally, Figure 2b shows that at large ; note that at large while at small , which leads to the cusp of each curve shown in the figure with its location (i.e., the -value at which ) increasing with increasing N (the cusp at N = 1 is located around ). We also note that , 0.119 and 0.182 for N = 1, 10 and 100. Therefore, with increasing , both and approach . Similar results are found for as shown in Figure 2c, where at large while at small (with the cusp at N = 1 located around ); also note that , 0.253 and 0.322 for N=1, 10 and 100. In particular, the PRISM-RPA theory with is equivalent to the Gaussian-fluctuation theory neglecting non-Gaussian fluctuations in the system and gives a correction to the SCF result, while the PRISM-PY theory captures non-Gaussian fluctuations in an approximate way and gives a leading-order correction to the Gaussian-fluctuation result. These are consistent with our previous study of compressible [36] and incompressible [37] homopolymer melts using fast lattice Monte Carlo simulations [6,7]. Given this and the agreement of our Figure 1b with experimental results at large
, we do not expect that the use of more accurate can qualitatively change our PRISM-PY results here.
To summarize, we have compared the correlation effects on the structural and thermodynamic properties of hard-core models (i.e., the THSC model and its limit of N→∞ at finite Re,0 (or equivalently at given
), the HC CGC-δ model [31]) and soft-core models (i.e., the DPD model and its limit of N→∞ at finite Re,0, the Edwards model [35]) for compressible homopolymer melts (or equivalently homopolymer solutions in an implicit solvent) given by the PRISM-PY theory. The behavior of soft-core models at large
depends on how
is varied, i.e., whether by changing N at fixed (thus ε is fixed) or by changing at fixed N (thus ε is also varied as being inversely proportional to ). In the former case, correlations exist even in the limit of N→∞, and both the hard-core and the DPD models give
at large
, consistent with real polymers used in experiments; it is, however, very difficult to reach via this way in molecular simulations even a relatively small
-value of 104 used in experiments. This problem is solved in the latter case, where the widely used polymer SCF theory becomes exact in the limit of (at finite N), the Gaussian-fluctuation theory gives a correction to the SCF result, and the PRISM-PY theory captures non-Gaussian fluctuations in the system in an approximate way and gives a leading-order correction to the Gaussian-fluctuation result, consistent with our previous simulations [36,37]. The soft-core models, however, give
at large
, suggesting that it would be difficult, if possible at all, for the various recently proposed simulation methods [3,4,5,6,7,8,9,10,11] to capture both the fluctuations and correlations in experimental systems. We also proposed an efficient numerical approach, which enables us to accurately solve the PRISM-PY theory for N as large as 106; numerical calculations of such theories can, therefore, capture both the fluctuations and correlations in experimental systems.
Supplementary Materials
The following supporting information can be downloaded at: https://www.mdpi.com/article/10.3390/polym15051180/s1, Figure S1: Logarithmic plot of the numerical errors given by pyPRISM and our approach for the HS model; Table S1: List of variables used in the main text.
Funding
This research and the APC were funded by the American Chemical Society Petroleum Research Fund (grant number 66122-ND6). Acknowledgment is made to the donors of The American Chemical Society Petroleum Research Fund for partial support of this research.
Data Availability Statement
The in-house code written in C and the data presented in this study are available upon reasonable request from the corresponding author.
Conflicts of Interest
The author declares no conflict of interest.
References
- Kremer, K.; Grest, G.S.; Carmesin, I. Crossover from Rouse to Reptation Dynamics: A Molecular-Dynamics Simulation. Phys. Rev. Lett. 1988, 61, 566–569. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Groot, R.D.; Warren, P.B. Dissipative particle dynamics: Bridging the gap between atomistic and mesoscopic simulation. J. Chem. Phys. 1997, 107, 4423–4435. [Google Scholar] [CrossRef] [Scilit]
- Laradji, M.; Guo, H.; Zuckermann, M.J. Off-Lattice Monte-Carlo Simulation of Polymer Brushes in Good Solvents. Phys. Rev. E 1994, 49, 3199–3206. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Wang, Q.; Yin, Y. Fast off-lattice Monte Carlo simulations with “soft” repulsive potentials. J. Chem. Phys. 2009, 130, 104903. [Google Scholar] [CrossRef] [Scilit]
- Detcheverry, F.A.; Pike, D.Q.; Nealey, P.F.; Muller, M.; de Pablo, J.J. Monte carlo simulation of coarse grain polymeric systems. Phys. Rev. Lett. 2009, 102, 197801. [Google Scholar] [CrossRef] [Scilit]
- Wang, Q. Studying soft matter with “soft” potentials: Fast lattice Monte Carlo simulations and corresponding lattice self-consistent field calculations. Soft Matter 2009, 5, 4564–4567. [Google Scholar] [CrossRef] [Scilit]
- Wang, Q. Correction-Studying soft matter with “soft” potentials: Fast lattice Monte Carlo simulations and corresponding lattice self-consistent field calculations. Soft Matter 2010, 6, 6206–6207. [Google Scholar]
- Fredrickson, G.H.; Ganesan, V.; Drolet, F. Field-theoretic computer simulation methods for polymers and complex fluids. Macromolecules 2002, 35, 16–39. [Google Scholar] [CrossRef] [Scilit]
- Matsen, M.W.; Beardsley, T.M. Field-Theoretic Simulations for Block Copolymer Melts Using the Partial Saddle-Point Approximation. Polymers 2021, 13, 2437. [Google Scholar] [CrossRef] [Scilit]
- Daoulas, K.C.; Muller, M. Single chain in mean field simulations: Quasi-instantaneous field approximation and quantitative comparison with Monte Carlo simulations. J. Chem. Phys. 2006, 125, 184904. [Google Scholar] [CrossRef] [Scilit]
- Milano, G.; Kawakatsu, T. Hybrid particle-field molecular dynamics simulations for dense polymer systems. J. Chem. Phys. 2009, 130, 214106. [Google Scholar] [CrossRef] [Scilit]
- Zong, J.; Wang, Q. Fluctuation/correlation effects in symmetric diblock copolymers: On the order-disorder transition. J. Chem. Phys. 2013, 139, 124907. [Google Scholar] [CrossRef] [Scilit]
- Fredrickson, G.H. The Equilibrium Theory of Inhomogeneous Polymers; Oxford University Press: New York, NY, USA, 2006. [Google Scholar]
- Yang, D.; Wang, Q. Systematic and simulation-free coarse graining of homopolymer melts: A structure-based study. J. Chem. Phys. 2015, 142, 054905. [Google Scholar] [CrossRef] [Scilit]
- Schweizer, K.S.; Curro, J.G. Integral-Equation Theory of the Structure of Polymer Melts. Phys. Rev. Lett. 1987, 58, 246–249. [Google Scholar] [CrossRef] [Scilit]
- Schweizer, K.S.; Curro, J.G. PRISM Theory of the Structure, Thermodynamics, and Phase-Transitions of Polymer Liquids and Alloys. Adv. Polym. Sci. 1994, 116, 319–377. [Google Scholar]
- Schweizer, K.S.; Curro, J.G. Integral equation theories of the structure, thermodynamics, and phase transitions of polymer fluids. Adv. Chem. Phys. 1997, 98, 1–142. [Google Scholar]
- Heine, D.R.; Grest, G.S.; Curro, J.G. Structure of polymer melts and blends: Comparison of integral equation theory and computer simulations. Adv. Polym. Sci. 2005, 173, 209–249. [Google Scholar]
- Martin, T.B.; Gartner, T.E.; Jones, R.L.; Snyder, C.R.; Jayaraman, A. pyPRISM: A Computational Tool for Liquid-State Theory Calculations of Macromolecular Materials. Macromolecules 2018, 51, 2906–2922. [Google Scholar] [CrossRef] [Scilit]
- Percus, J.K.; Yevick, G.J. Analysis of Classical Statistical Mechanics by Means of Collective Coordinates. Phys. Rev. 1958, 110, 1–13. [Google Scholar] [CrossRef] [Scilit]
- Lowden, L.J.; Chandler, D. Solution of a new integral equation for pair correlation functions in molecular liquids. J. Chem. Phys. 1973, 59, 6587–6595. [Google Scholar] [CrossRef] [Scilit]
- Yethiraj, A.; Schweizer, K.S. Self-consistent polymer integral equation theory: Comparisons with Monte Carlo simulations and alternative closure approximations. J. Chem. Phys. 1992, 97, 1455–1464. [Google Scholar] [CrossRef] [Scilit]
- Talitskikh, S.K.; Khalatur, P.G. Solvent Effect on Conformations of Hydrocarbon Molecules: Model of Atom-Atom Potentials and Integral-Equations of Liquid Theory. Zhurnal Fizicheskoi Khimii 1995, 69, 5–11. [Google Scholar]
- Heine, D.; Wu, D.T.; Curro, J.G.; Grest, G.S. Role of intramolecular energy on polyolefin miscibility: Isotactic polypropylene/polyethylene blends. J. Chem. Phys. 2003, 118, 914–924. [Google Scholar] [CrossRef] [Scilit]
- Britanak, V.; Yip, P.C.; Rao, K.R. Discrete Cosine and Sine Transforms: General Properties, Fast Algorithms and Integer Approximations; Academic: Amsterdam, The Netherlands; Boston, MA, USA, 2007. [Google Scholar]
- Zhang, P.; Wang, Q. Repulsion between Colloidal Particles Mediated by Nonadsorbing Polymers: Lattice Monte Carlo Simulations and the Corresponding Self-Consistent Field Calculations. Macromolecules 2019, 52, 5777–5790. [Google Scholar] [CrossRef] [Scilit]
- Press, W.H. Chapter 4.3. In Numerical Recipes in C: The Art of Scientific Computing, 2nd ed.; Cambridge University Press: Cambridge, UK; New York, NY, USA, 1992. [Google Scholar]
- Matsen, M.W. Fast and accurate SCFT calculations for periodic block-copolymer morphologies using the spectral method with Anderson mixing. Eur. Phys. J. E 2009, 30, 361–369. [Google Scholar] [CrossRef] [Scilit]
- de Gennes, P.G. Scaling Concepts in Polymer Physics; Cornell University Press: Ithaca, NY, USA, 1979. [Google Scholar]
- Ornstein, L.S.; Zernike, F. Acculental deviations of density and opalescence at the critical point of a simple substance. Proc. K. Akad. Wet.-Amst. 1914, 17, 793–806. [Google Scholar]
- Schweizer, K.S.; Curro, J.G. RISM Theory of Polymer Liquids: Analytical Results for Continuum Models of Melts and Alloys. Chem. Phys. 1990, 149, 105–127. [Google Scholar] [CrossRef] [Scilit]
- Fredrickson, G.H.; Helfand, E.; Bates, F.S.; Leibler, L. Microphase Separation in Block Copolymers-Recent Developments. Springer Ser. Chem. Phys. 1989, 51, 13–19. [Google Scholar]
- Lang, A.; Likos, C.N.; Watzlawek, M.; Lowen, H. Fluid and solid phases of the Gaussian core model. J. Phys. Condens. Matter 2000, 12, 5087–5108. [Google Scholar] [CrossRef] [Scilit]
- Louis, A.A.; Bolhuis, P.G.; Hansen, J.P. Mean-field fluid behavior of the Gaussian core model. Phys. Rev. E 2000, 62, 7961–7972. [Google Scholar] [CrossRef] [Scilit]
- Edwards, S.F. The theory of polymer solutions at intermediate concentration. Proc. Phys. Soc. 1966, 88, 265–280. [Google Scholar] [CrossRef] [Scilit]
- Zhang, P.; Zhang, X.; Li, B.; Wang, Q. Quantitative study of fluctuation effects by fast lattice Monte Carlo simulations. I. Compressible homopolymer melts. Soft Matter 2011, 7, 4461–4471. [Google Scholar] [CrossRef] [Scilit]
- Zhang, P.; Yang, D.; Wang, Q. Quantitative study of fluctuation effects by fast lattice Monte Carlo simulations. V. Incompressible homopolymer melts. J. Phys. Chem. B 2014, 118, 12059–12067. [Google Scholar] [CrossRef] [Scilit]
Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2023 by the author. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license (https://creativecommons.org/licenses/by/4.0/).