


Abstract
The segregation of niobium (Nb) at the FeΣ3(111) grain boundary and its influence on interfacial cohesion were investigated via spin-polarized density functional theory calculations. Nb atoms exhibit strong site-selective segregation, with Site 1 being the most thermodynamically favorable one (segregation energy of −2.47 eV), owing to its largest local Voronoi volume. Electronic structure analyses reveal pronounced Nb-4d/Fe-3d orbital hybridization and localized charge accumulation between Nb and neighboring Fe atoms, enhancing covalent bonding at the boundary. First-principles tensile simulations show that single-Nb segregation increases the critical strain from 13.58% to 15.76% and the theoretical tensile strength from 16.32 GPa to 19.64 GPa. However, double-Nb segregation reduces the work of separation to 3.26–4.24 J/m2, revealing a competition between segregation strengthening and solute-induced weakening that implies an optimal Nb concentration window for grain boundary engineering.
1. Introduction
As the cornerstone of modern industrial civilization, steel plays an irreplaceable strategic role in national economic development and national defense. Among the various steel material systems, stainless steel has emerged as an indispensable key material in high-end manufacturing fields such as aerospace, petrochemicals, energy equipment, transportation, and architectural decoration due to its outstanding corrosion resistance, excellent mechanical properties, and superior surface decorative properties [1,2]. Based on the characteristics of their room-temperature crystal structure, stainless steels can be classified into five major categories, namely austenitic stainless steel, ferritic stainless steel, duplex stainless steel, martensitic stainless steel, and precipitation-hardening stainless steel. Among them, ferritic stainless steel, due to its body-centered cubic crystal structure, possesses unique physical metallurgical properties and demonstrates unparalleled application advantages in specific service environments.
As the most critical microalloying element in 442D stainless steel, Nb is typically present at contents of about 0.3–0.6%, existing predominantly in two forms—solid solution and Nb(C,N) precipitates—thereby exerting a decisive influence on the microstructural evolution and property regulation of the material. During the hot processing and heat treatment, the diffusion, segregation, and precipitation behavior of Nb atoms directly determine the grain size, texture, dislocation density, and interface characteristics, thereby profoundly influencing the mechanical properties, corrosion resistance, and manufacturability [3,4]. However, the physical metallurgy of Nb in ferritic stainless steel is extremely complex, involving coupling across multiple length scales and physical fields. At the atomic scale, Nb atoms tend to segregate at defect sites and interfaces, thereby influencing the stability of the interface by altering the local electronic structure and atomic bonding characteristics. At the nanoscale, Nb combines with C and N atoms to form Nb(C,N) precipitates, which contribute to precipitation strengthening and grain boundary pinning. At the micro scale, the solute drag and the pinning effect of Nb-containing precipitates interact to influence recrystallization kinetics and grain growth [5,6]. A thorough understanding of the multi-scale precipitation behavior of Nb from the atomic scale to the micro scale and its influence mechanism on the microstructures and properties has significant theoretical value and engineering importance for optimizing the heat treatment of 442D ferritic stainless steel and achieving precise control of microstructures and properties. Recent progress in computational materials science has enabled systematic first-principles investigations of transition metal segregation at ferritic iron grain boundaries. Mai et al. [7] performed a high-throughput DFT study of 10 transition metals (including Nb) with various grain boundary characters in α-Fe, demonstrating that segregation energies and cohesion effects vary significantly with the boundary type and that solute–solute interactions at boundaries differ substantially from those in the bulk. These findings highlight the necessity of characterizing site-specific segregation behavior at well-defined grain boundary models.
In BCC-structured low-alloy steels, grain boundaries not only impede dislocation motion but also serve as the most preferential sites for crack initiation. Researchers have found that a trace amount of Nb (<0.6 wt.%) can significantly enhance the grain boundary cohesion strength of steel and mitigate intergranular fracture tendencies [8]. However, a systematic and quantitative atomic-scale understanding of Nb segregate at grain boundaries and its electronic-level strengthening mechanism remains lacking. The FeΣ3(111) symmetric tilt grain boundary belongs to the coincidence site lattice (CSL) model with a low Σ value (Σ = 3) and is one of the most common and lowest-energy high-angle grain boundaries in bcc-Fe. CSL boundaries are characterized by a periodic arrangement of atoms where a fraction 1/Σ of the lattice sites coincide across the interface, giving them special thermodynamic stability. Its periodic structure facilitates the construction of computationally tractable supercells. Experimental results indicate that recrystallization of the material is largely completed after heat treatment at 900 °C [9]. The grain boundary misorientation distribution exhibits a pronounced peak in the range of 50–60°, which is close to the theoretical misorientation of 60° for the Σ3(111) twin boundary in the BCC structure, suggesting that a large number of low-energy special grain boundaries are formed during recrystallization. The proportion and distribution of these Σ3 twin boundaries directly affect the mechanical properties of the material. At 900 °C, the material achieves optimal plasticity, and the fracture mode transitions from cleavage to dimple rupture. However, there is a lack of direct atomic-scale evidence on how a small amount of Nb (0.51 wt.%) achieves strengthening through segregating at these twin boundaries. In this work, an FeΣ3(111) symmetric tilt grain boundary model is constructed to systematically investigate the site preference of Nb atoms, the electronic structure characteristics, and the influence of Nb segregation on grain boundary strength, as well as the microscopic mechanisms underlying the mechanical properties. To elucidate the electronic-level strengthening mechanism, two key quantum-mechanical analyses will be employed. The density of states (DOS) describes the distribution of electronic states as a function of energy and enables the identification of orbital hybridization between the segregated solutes and host Fe atoms. The charge density difference visualizes the spatial redistribution of electrons upon solute segregation, providing direct evidence for the formation of directional covalent bonds and local charge transfer. These electronic structure tools are essential for bridging atomic segregation behavior with macroscopic grain boundary cohesion.
It should be emphasized that the present study focuses exclusively on the symmetric FeΣ3(111)[1−10] tilt grain boundary, which represents a special low-energy boundary in body-centered cubic iron. While this boundary is frequently observed in recrystallized ferritic stainless steels and serves as an ideal prototype for atomistic simulations, owing to its periodic structure, the conclusions drawn herein should not be uncritically generalized to random high-angle grain boundaries or to the full heterogeneous grain boundary network present in engineering steels. The Σ3(111) boundary is selected as a necessary first step to establish the intrinsic electronic and mechanical response of Nb segregation at a well-defined interface [10,11], providing a baseline for future comparative studies involving Σ5, Σ7, and other general grain boundaries.
Despite the recognized importance of Nb in ferritic stainless steels, a systematic atomic-scale understanding of its site-selective segregation at Σ3 twin boundaries and the associated electronic-level strengthening mechanism remains lacking. In particular, three questions are unresolved. (1) Which specific sites at the FeΣ3(111) boundary are thermodynamically preferred by Nb? (2) How does the local electronic structure (orbital hybridization and charge redistribution) vary by segregation site? (3) Finally, at what concentration does the beneficial cohesion enhancement transition to solute-induced weakening? This work employs spin-polarized density functional theory (DFT) calculations to address these questions, aiming to provide an atomistic rationale that is qualitatively compatible with reported macroscopic behavior at 900 °C while acknowledging that direct experimental validation of the calculated configurations requires future atom probe tomography (APT) or high-resolution transmission electron microscopy (HRTEM) investigations.
2. Calculation Details and Model Construction
2.1. Computational Methods and Details
All calculation work presented in this chapter was performed using the CASTEP (version 2020) software package, which is based on density functional theory combined with the plane wave pseudopotential method [12]. For bulk bcc-Fe, structural optimization used a Monkhorst-Pack k-point grid of 4 × 4 × 4. The GGA-PBE exchange correlation functional for electrons was selected, and the Monkhorst-Pack method was used to divide the Brillouin zone into grids [13]. To ensure the reliability of the calculated energies, systematic convergence tests were performed. The plane-wave cutoff energy was tested from 300 eV to 500 eV in increments of 50 eV; the results indicate that the energy began to converge when the cutoff exceeded 400 eV. The k-point mesh was tested from 3 × 2 × 1 to 7 × 5 × 1, and the energy began to converge at a mesh of 5 × 3 × 1, which was therefore adopted. All calculations were performed with spin polarization using a ferromagnetic ground state, which is appropriate for low-alloy bcc-Fe at the dilute Nb concentrations considered here. A k-point grid of 5 × 3 × 1 was employed to sample the Brillouin zone. The plane-wave cutoff energy was set to 400 eV. During structural optimization, the constructed model contained atomic layers in the vacuum region. The outermost 4 layers of atoms were fixed to keep the cell shape unchanged, while the remaining atoms were allowed to relax fully until convergence. Convergence criteria were set to a total energy change below 10−6 eV/atom and residual forces below 1 × 10⁻⁴ eV/Å.
2.2. The Lattice Constant of Bcc-Fe
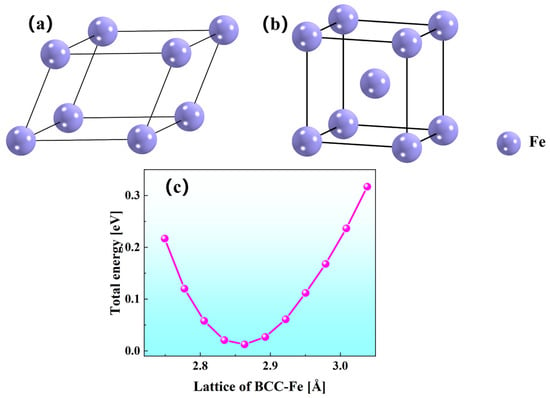
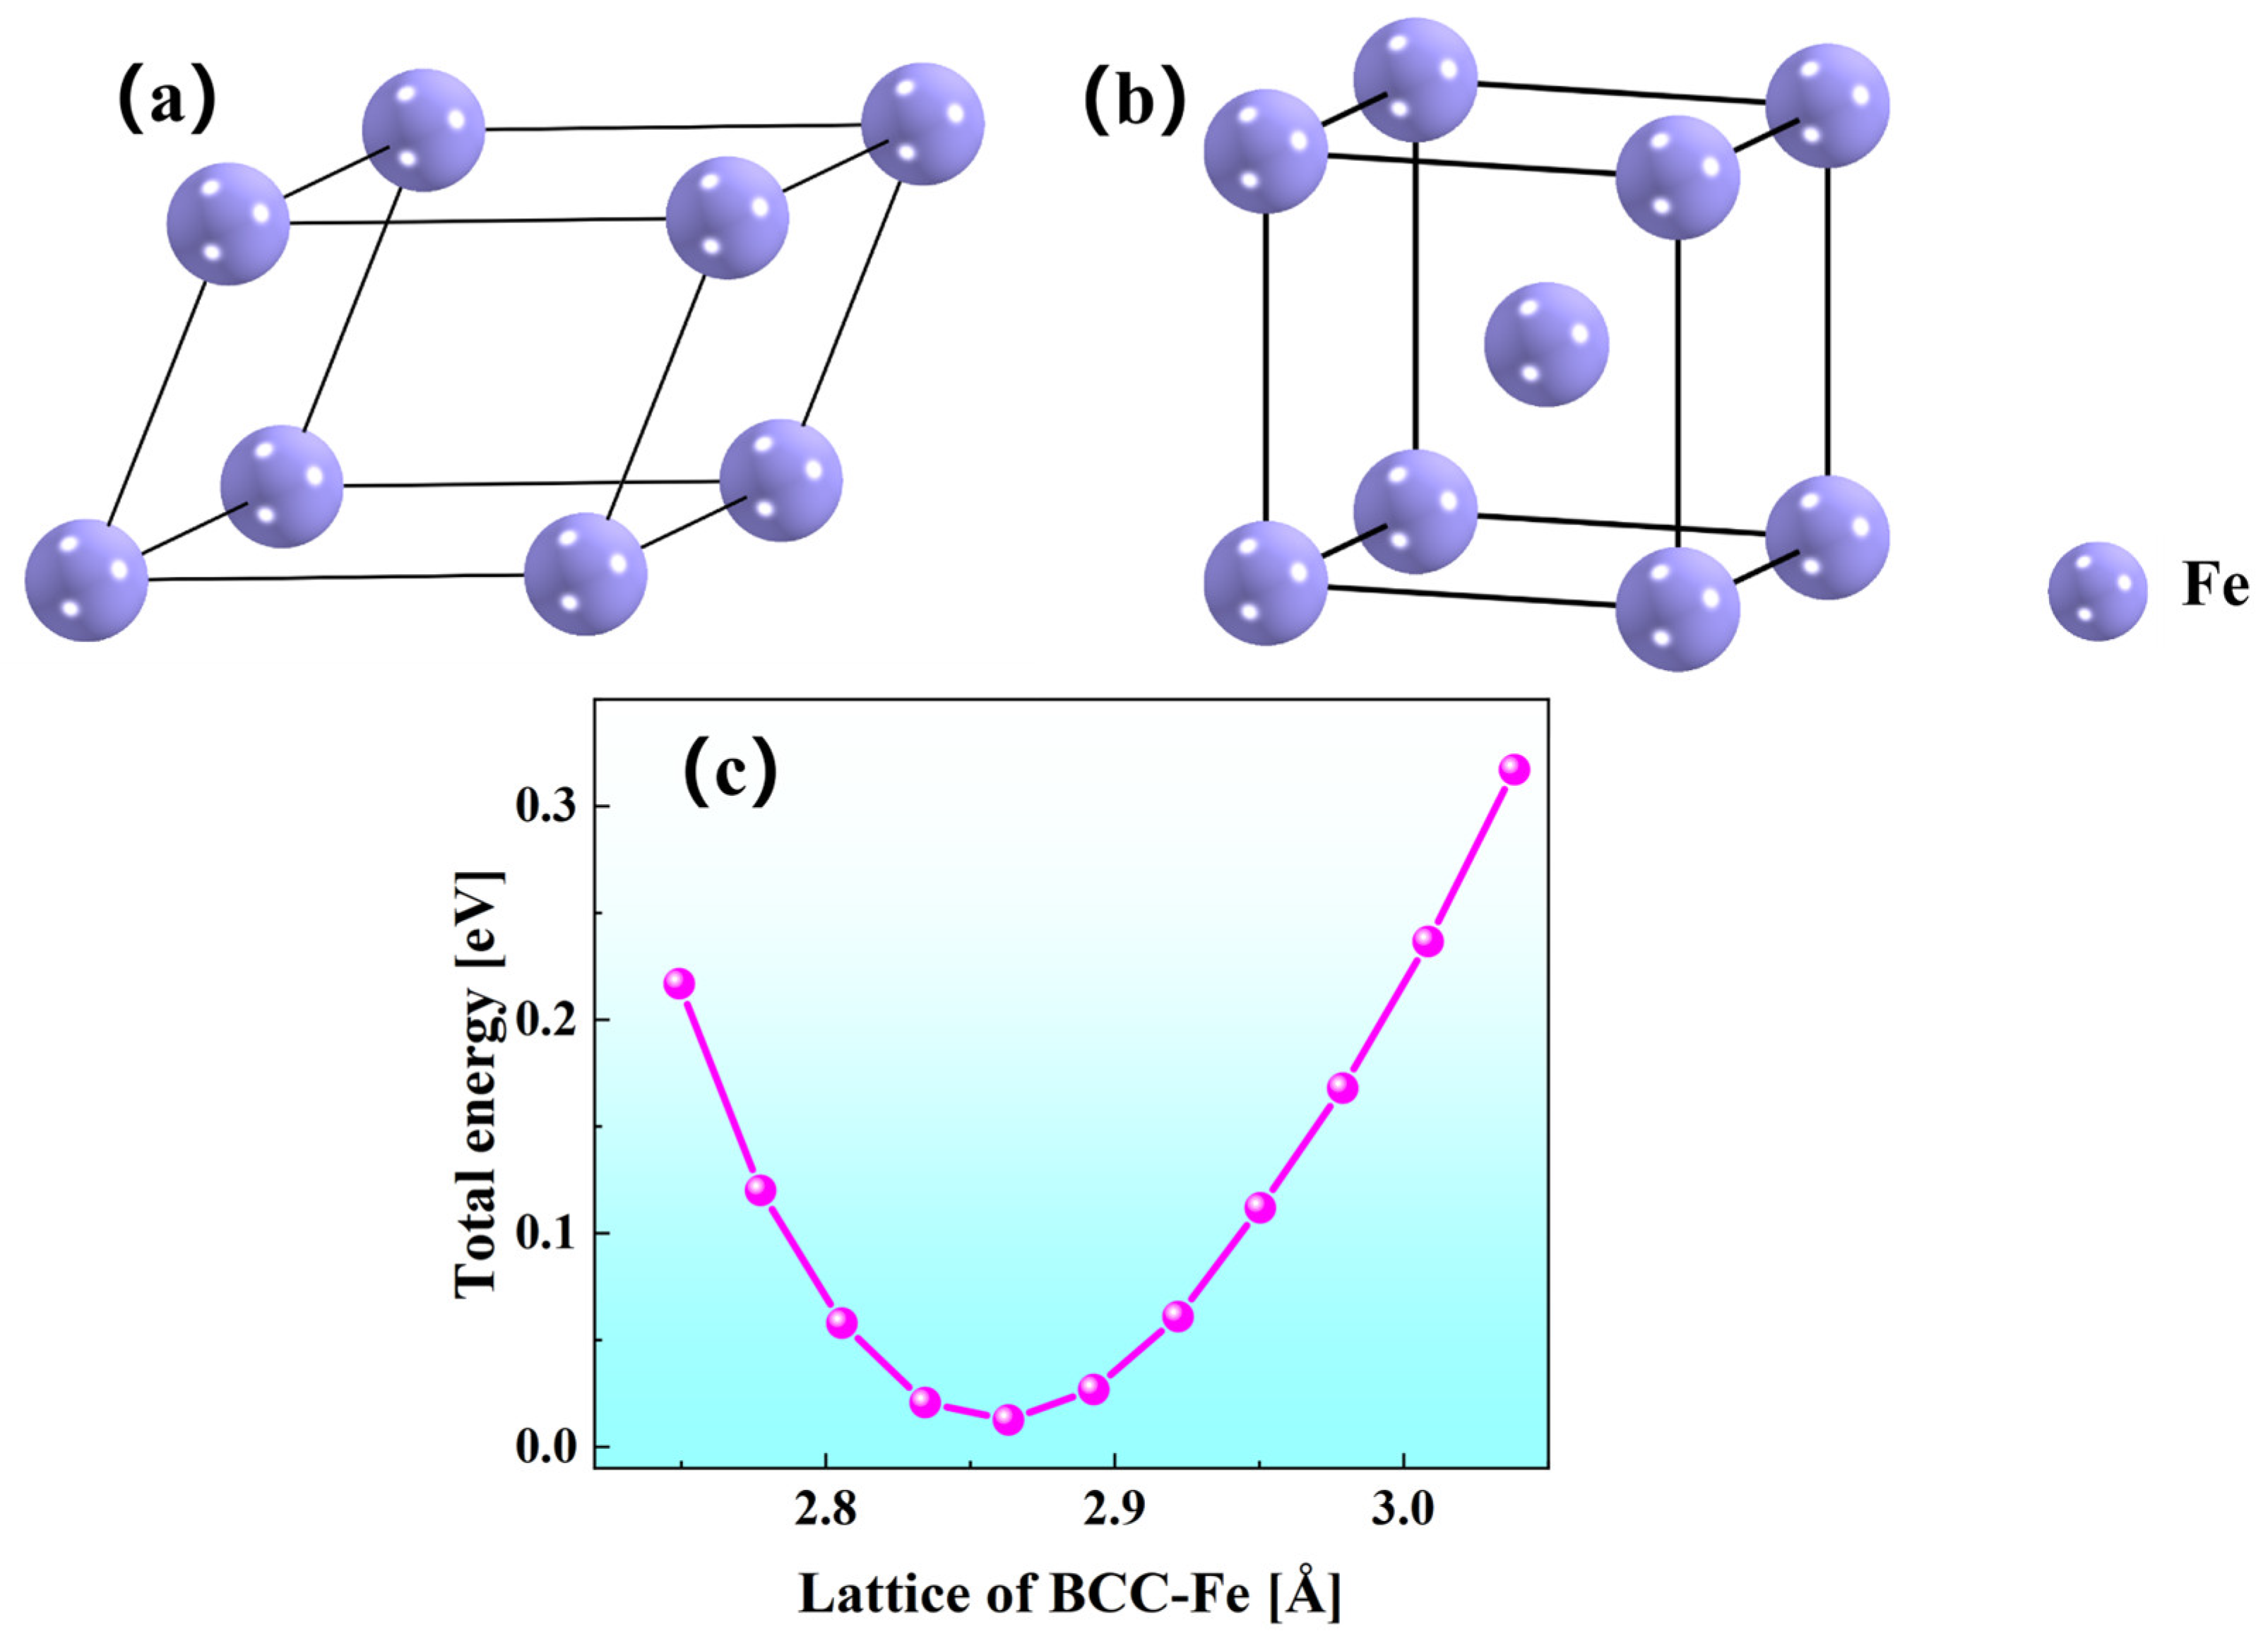
To validate the reliability of the pseudopotentials and computational parameters employed, first-principles calculations were performed to determine the precise lattice constant of bcc-Fe. Figure 1 illustrates the crystal structure model of bcc-Fe. Given the magnetic nature of Fe, all calculations were carried out with spin polarization. By varying the lattice constant of Fe incrementally, the total energy of the system under the corresponding conditions was calculated. According to the principles of thermodynamics, the lower the total energy of the system, the stronger its thermodynamic stability. Figure 1 presents the relationship between the lattice constant of Fe and the total energy of the system. After calculation, the optimal lattice constant of Fe was 2.863 Å. This was compared with the experimental value of 2.866 Å reported in the literature [14], with an error of only 0.1%. The close agreement between the theoretical and experimental lattice constants confirms the reliability of the selected Fe pseudopotential and validates the computational parameters employed.
Figure 1.
The crystal structure model of bcc-Fe: (a) unit cell and (b) primitive cell. (c) The relationship curve between the lattice constant of bcc-Fe and the total energy of the system.
2.3. Construction of the FeΣ3(111) Grain Boundary Model
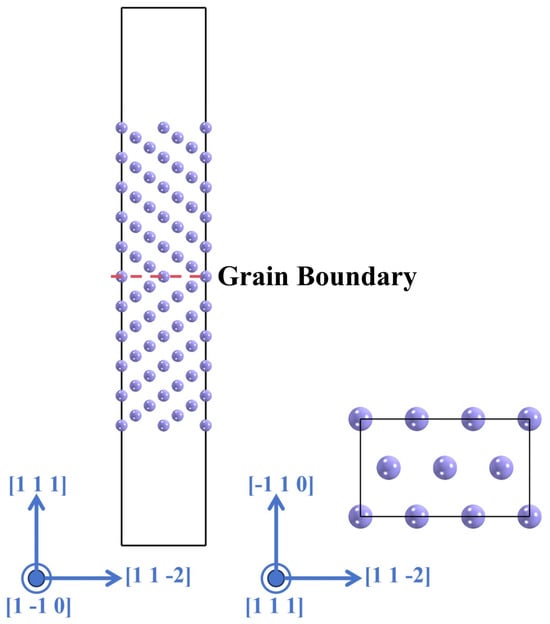
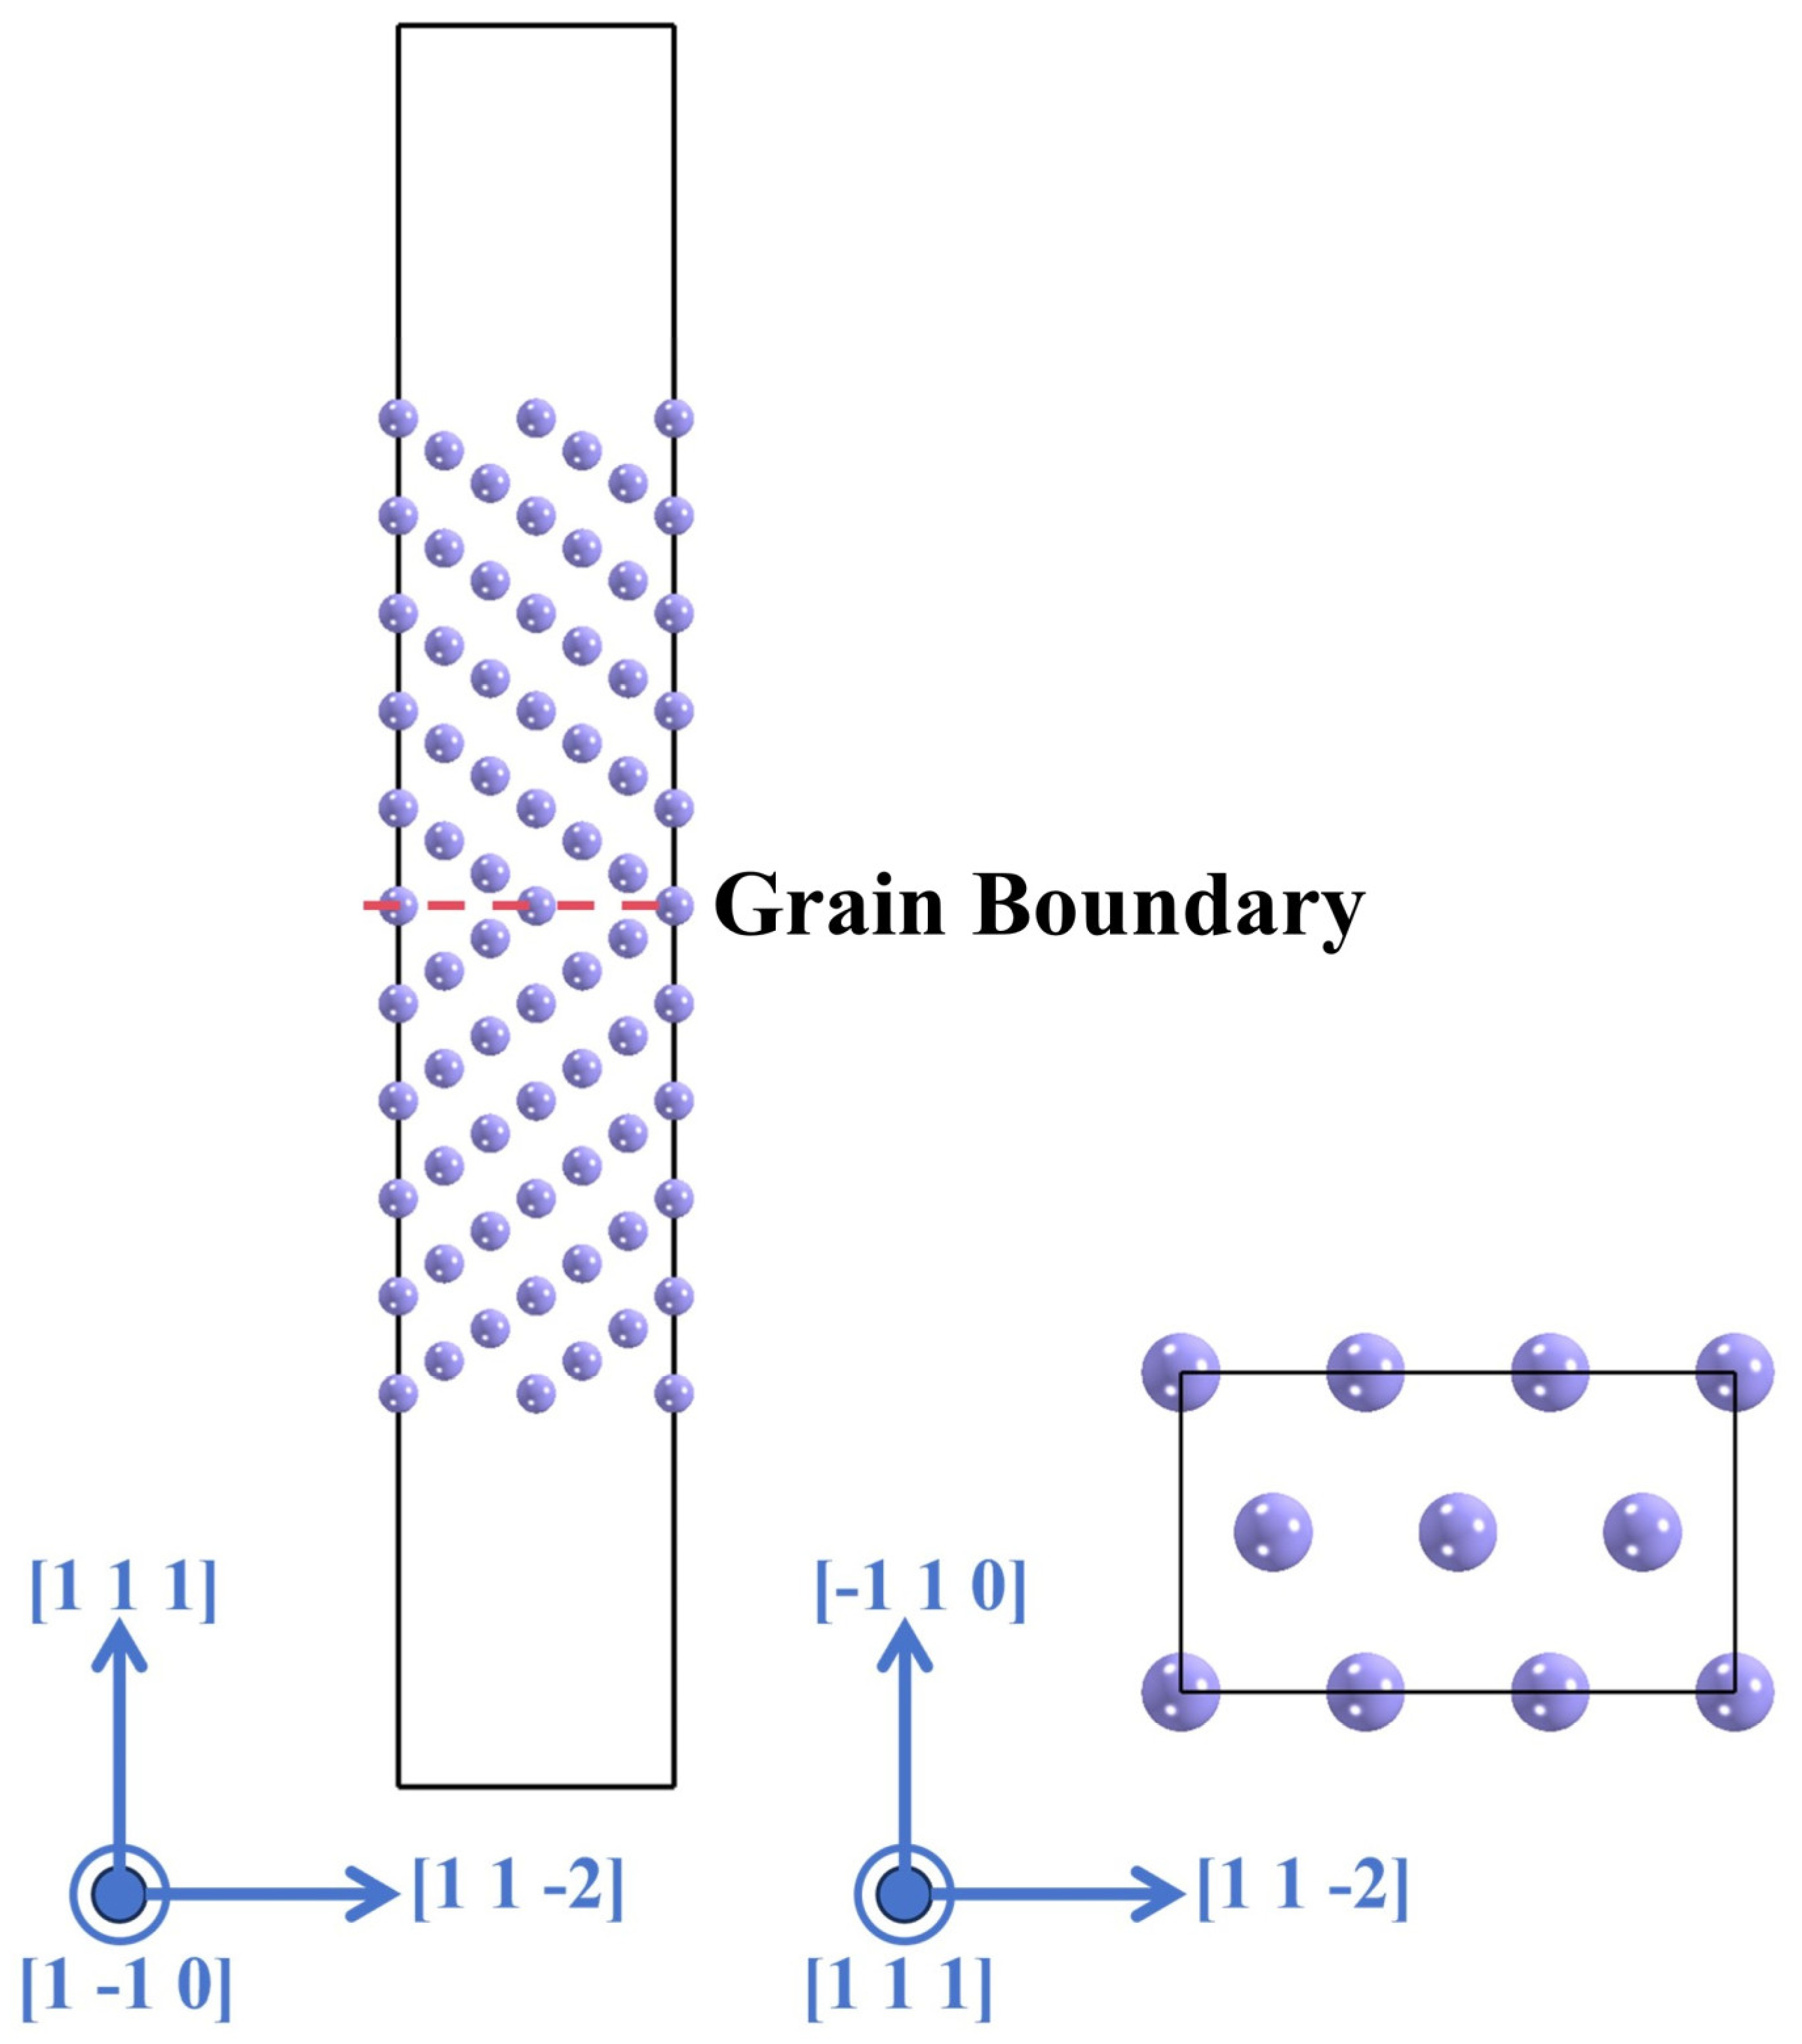
In this work, a symmetrically tilted FeΣ3(111)[1−10] grain boundary was selected as the research object. As a typical high-angle grain boundary that can exist stably in bcc-Fe, it is suitable for investigating the effects of Nb segregation on the FeΣ3 grain boundary and its mechanical properties. FeΣ3(111)[1−10] is a notation used to describe a grain boundary in a crystal, where Σ3 indicates that one-third of the lattice points on either side of the boundary coincide and [1−10] and (111) represent the Miller indices of the tilt axis and the grain boundary plane, respectively. Figure 2 presents the symmetrically tilted FeΣ3(111)[1−10] of the grain boundary model. Among them, the x, y, and z axes were oriented along [111], [11−2], and [1−10] directions, respectively, and 15 atomic layers were stacked along the [111] direction, while vacuum layers of 10 Å were added on both sides to avoid the interaction between periodic images. This model was composed of 62 atoms, and the optimized size of the grain boundary supercell was 45.26 × 3.94 × 6.91 Å3.
Figure 2.
Model of FeΣ3(111)[1−10] symmetrically inclined grain boundary.
The grain boundary energy (γGB) is a key physical quantity that characterizes the stability of grain boundaries. It is defined as the change in system energy per unit area of interface formed, which is numerically equal to the energy per unit area of the interfacial region minus the energy per unit area of the same region without the interface. The specific formula for calculating the grain boundary energy is as follows:
Here, is the total energy of the grain boundary, is the energy of a single Fe atom in a perfect Fe unit cell, is the number of Fe atoms in the grain boundary model under study, and is the cross-sectional area of the grain boundary.
Table 1 presents the calculated grain boundary energy for the symmetric FeΣ3(111)[1−10] tilt boundary, along with corresponding results reported by other researchers. The value of 1.60 J/m2 obtained in this work agrees well with the literature data, indicating that the model constructed and the parameter settings used in the calculation are reasonable.
Table 1.
Crystalline boundary energy of FeΣ3(111)[1−10] symmetric inclined boundaries.
3. Results and Discussion
3.1. The Occupation Tendency of Nb at the FeΣ3(111) Grain Boundary
Clarifying the preference of solute atoms to occupy specific positions at the FeΣ3(111) grain boundary is essential for revealing the effect of grain boundary segregation. Whether Nb atoms tend to accumulate near grain boundaries or remain dispersed in the matrix can be determined by evaluating the thermodynamic driving force for solute segregation to the boundary via theoretical calculations.
From the perspective of theoretical calculation, the site preference of atoms can be explained by comparing the stability of the system’s energy. Therefore, the segregation energy () is introduced. It can be defined as the energy difference between a system with segregated elements and a reference system without segregation. The sign and magnitude of this energy difference reflect the segregation tendency of the solute atom at the grain boundary. The formula is as follows [16]:
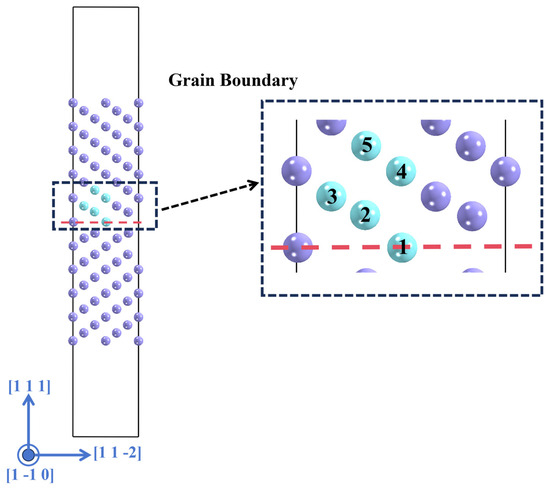
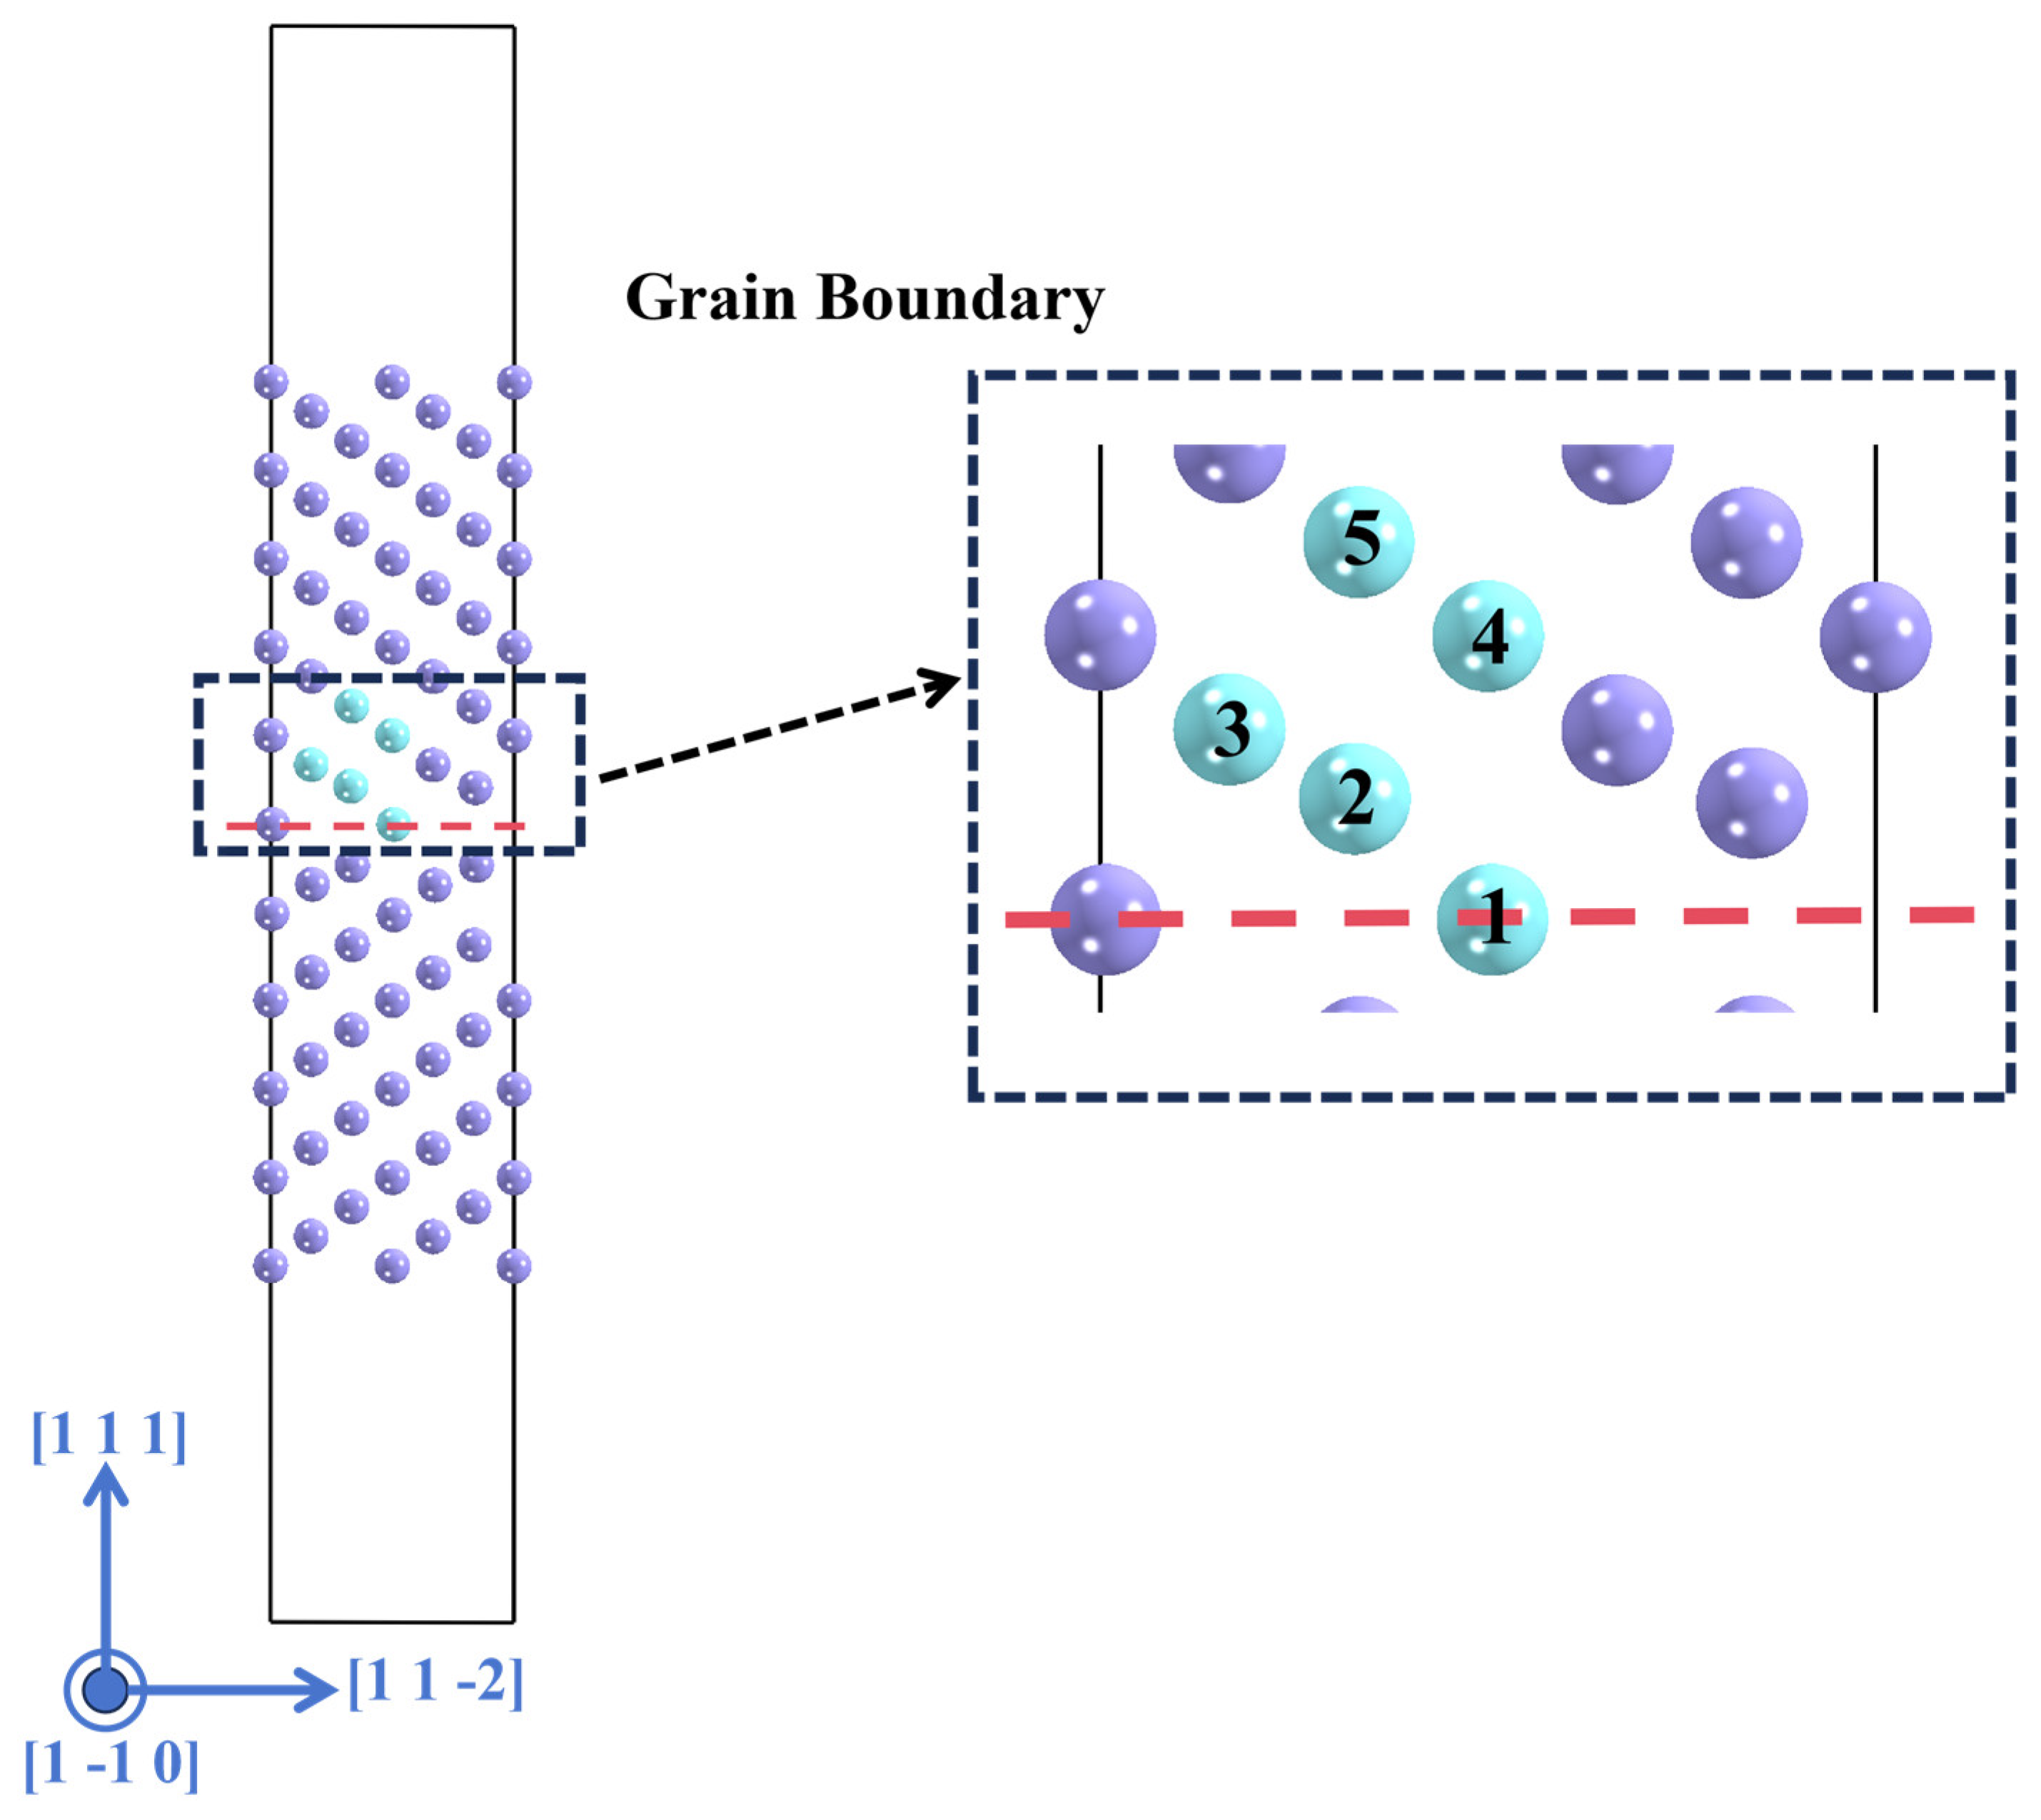
Here, and represent the total energy of the system with one solute atom A located at the grain boundary and in the bulk, respectively; represents the total energy of a clean Fe grain boundary; and Ebulk represents the total energy of the Fe bulk. The structures of and both consist of a 30-layer atomic Fe bulk layer crystal model containing 62 atoms. As can be seen from Equation (2), a negative value of Eseg indicates that the atom tends to segregate toward the grain boundary, while a positive value of Eseg suggests that it remains more stable in the matrix. The methods of solute segregation can be classified into two types: substitutional segregation and interstitial segregation. The covalent radius of Nb atoms was 0.146 nm, while that of Fe atoms was 0.140 nm. Since the radius of Nb atoms was larger than that of Fe, this resulted in a positive size mismatch. Therefore, from a thermodynamic perspective, Nb atoms are therefore expected to have a strong driving force to segregate to Fe grain boundaries, thereby reducing the overall system energy. The Nb atoms replace the occupied positions on the grain boundaries, as shown in Figure 3. The five initial sites in Figure 3 were preliminarily identified based on the symmetrically asymmetric positions along crystal boundaries and Voronoi volumes.
Figure 3.
Five segregation sites at the FeΣ3(111) grain boundary.
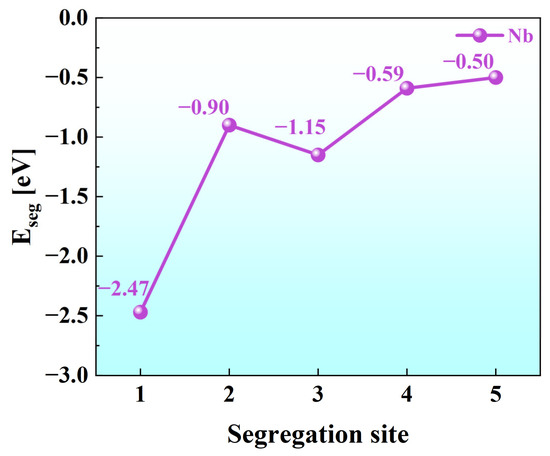
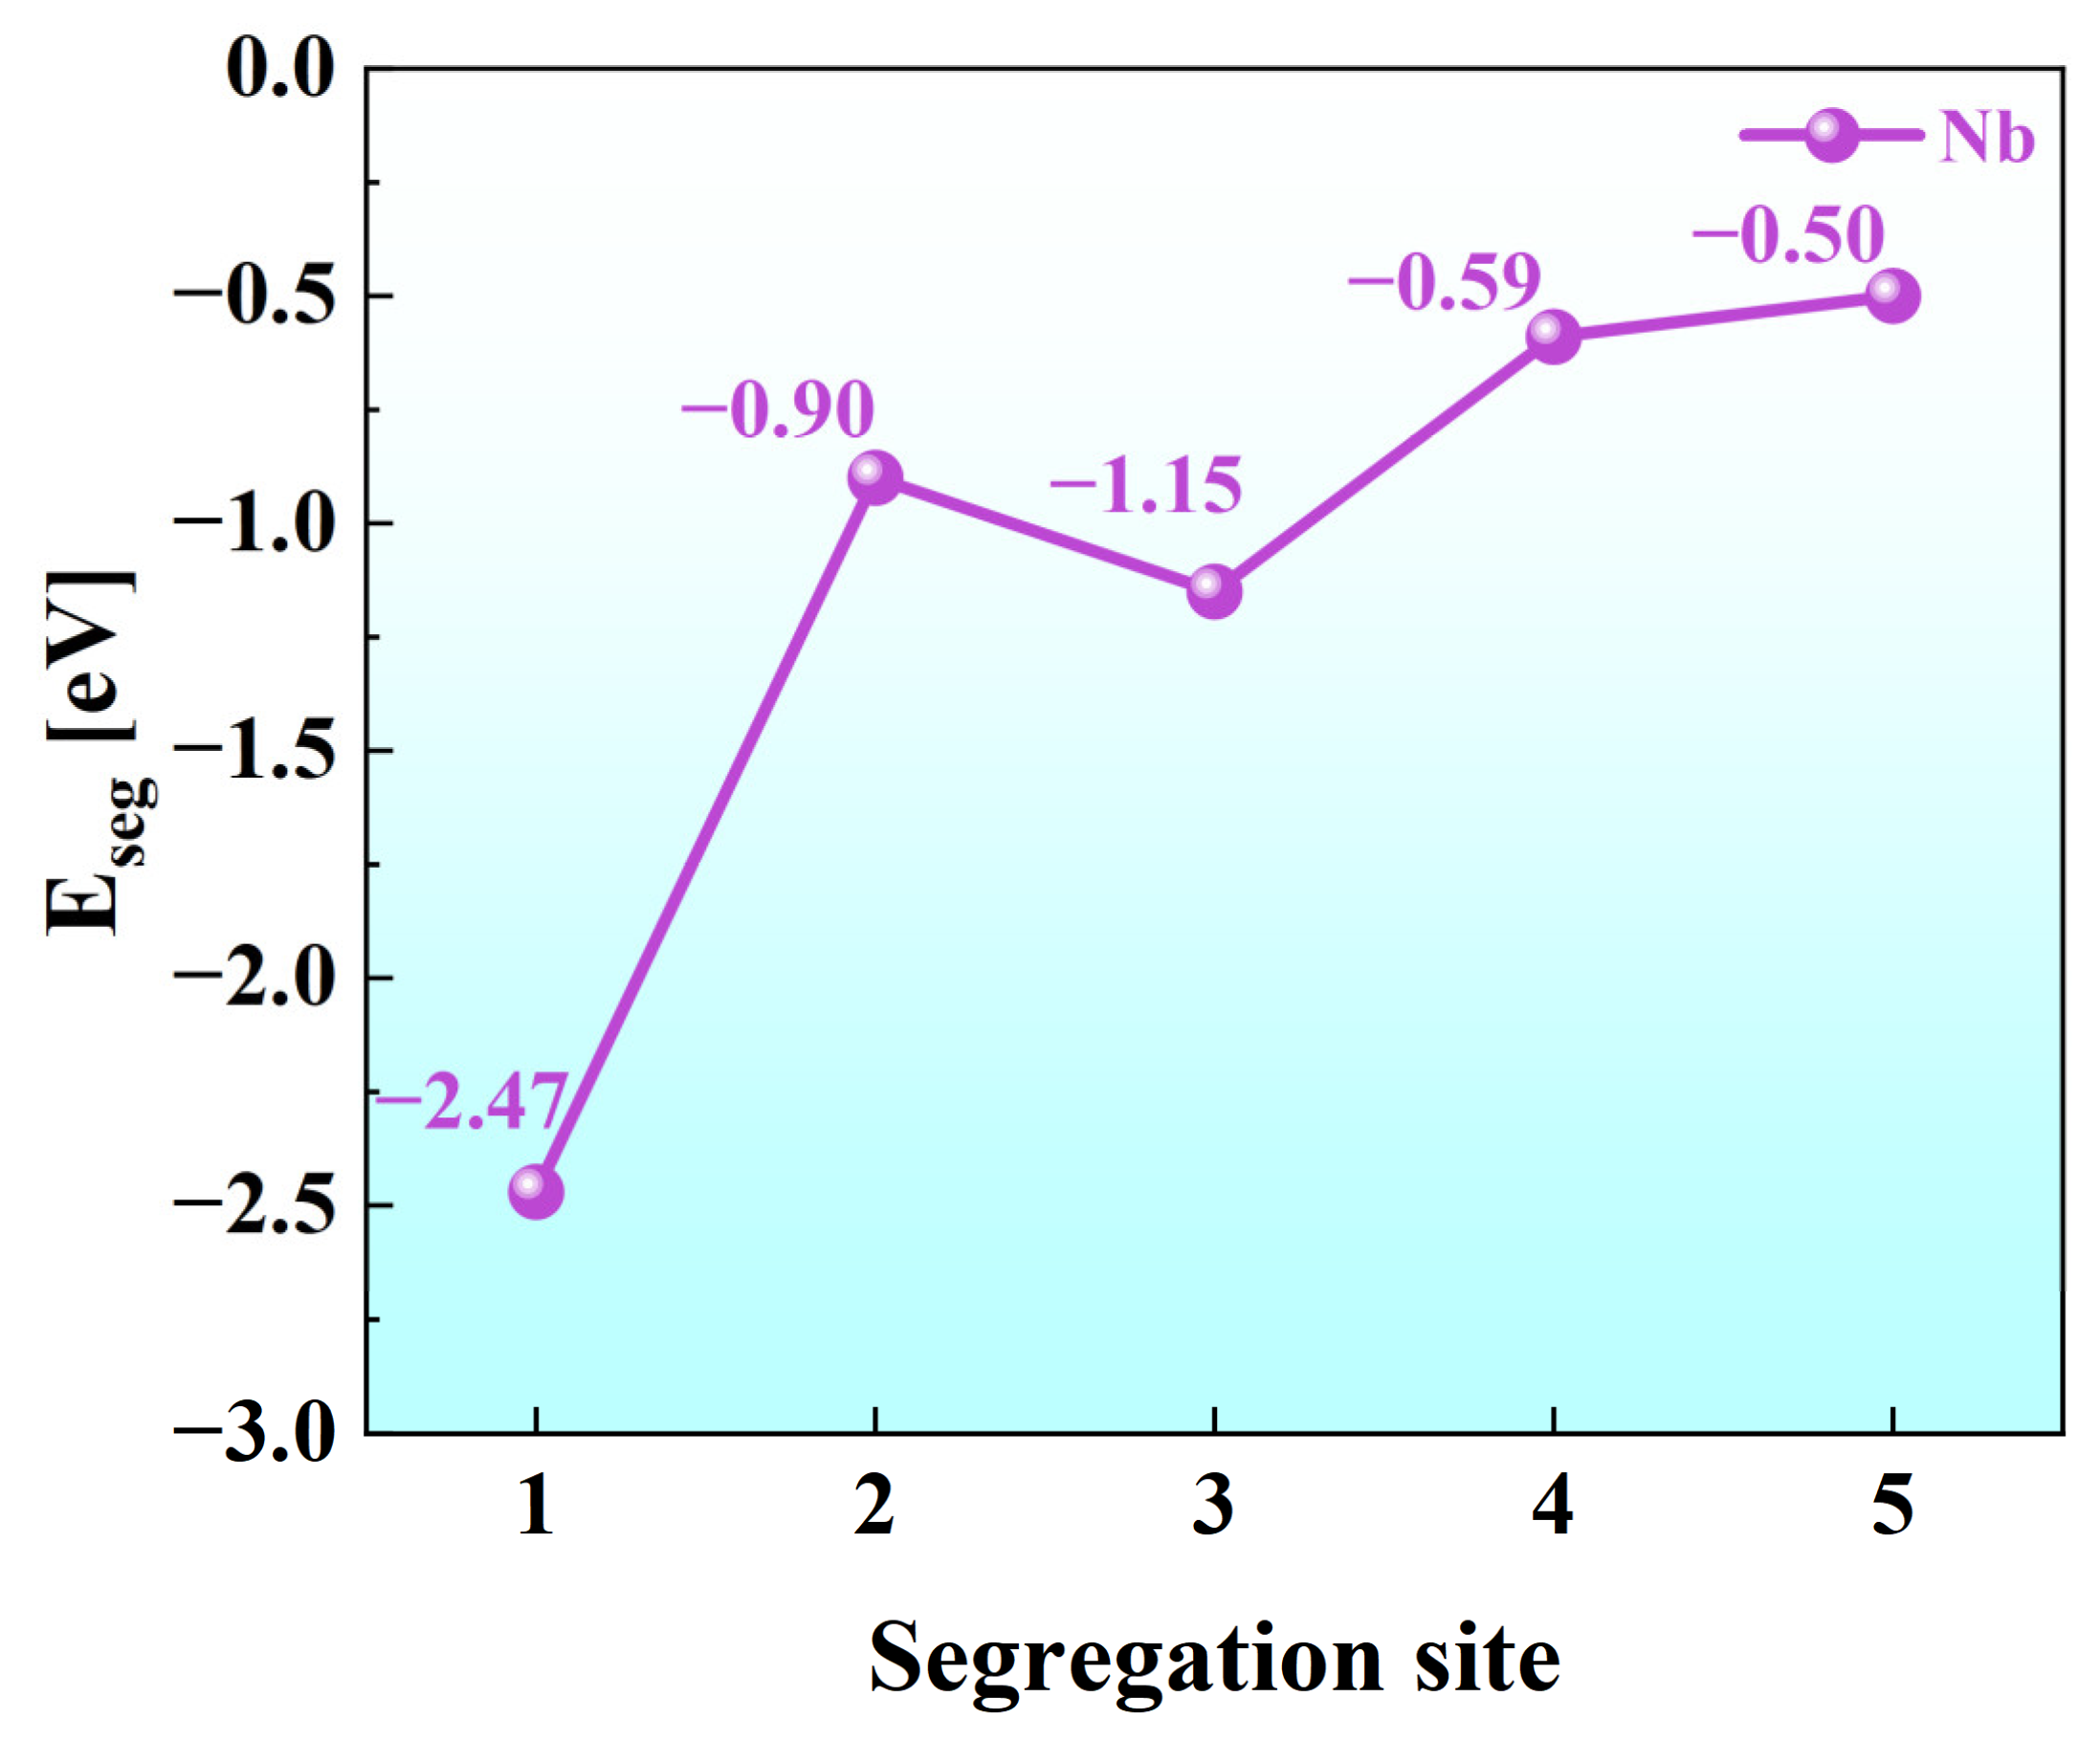
The calculated results of the segregation energy of Nb atoms at the above five grain boundary sites are shown in Figure 4. It is clear that there were significant differences in the segregation energy of Nb atoms at different sites. The overall trend shows that from site 1 to site 5, the value of the segregation energy increased. Among them, site 1 exhibited the lowest segregation energy of −2.47 eV, and the absolute value of its negative value was much larger than that of the other sites. This indicates that the Nb atoms exhibited strong site selectivity at the Σ3(111) grain boundary, and they were most likely to occupy site 1. In contrast, the negative values of the segregation energy at sites 4 and 5 were smaller, indicating a weaker segregation tendency and lower stability at these two locations.
Figure 4.
The segregation energy of Nb atoms at different FeΣ3(111) grain boundary sites.
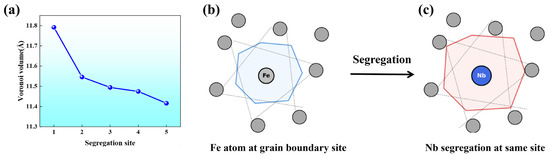
Figure 5a presents the Voronoi volumes of five segregated sites at the FeΣ3(111) grain boundary. To clarify the physical meaning of the Voronoi volume, a schematic illustration is provided in Figure 5b,c. The Voronoi polyhedron of an atom is constructed from the perpendicular bisector planes to its neighboring atoms, enclosing the region of space closer to that atom than to any other. Because the covalent radius of Nb (0.146 nm) was larger than that of Fe (0.140 nm), substitutional segregation induced local lattice expansion. Sites with larger initial free volumes can better accommodate this oversized solute, minimizing the elastic strain energy and making segregation thermodynamically favorable. From site 1 to site 5, the corresponding Voronoi volume continued to decrease. Site 1 had the largest volume, while Site 5 showed the smallest, indicating that the atomic arrangement at site 1 was the most loosely packed one, providing the greatest space to accommodate foreign atoms. This volume trend shows a clear negative correlation with the Nb segregation energies presented in Figure 4. In comparison, at site 1, which had the largest volume, the segregation energy of the Nb atom was also the most negative one, reaching −2.47 eV, indicating that Nb atoms most readily segregated to this site and formed the most stable bonding there. In contrast, as the site volume decreased toward sites 4 and 5, the negative values of the segregation energy of Nb atoms at these sites also decreased significantly, reaching −0.59 eV and −0.50 eV, respectively, reflecting a much weaker segregation tendency and lower stability. The results indicate that in the FeΣ3(111) grain boundary, the local spatial volume size of the sites was one of the key factors influencing the Nb atom segregation behavior. Larger-volume sites can more effectively accommodate the larger-sized Nb atoms, thereby reducing the lattice strain energy and making the segregation process more energetically favorable. The larger Voronoi volumes observed at the grain boundary sites reflect the inherently lower atomic coordination number and the presence of excess free volume at interfaces compared with the perfect bulk lattice. This is a general characteristic of crystalline grain boundaries. However, the magnitude of the excess volume depends strongly on the boundary structure. For the low-energy Σ3(111) CSL twin boundary investigated here, the excess volume was relatively modest because of its highly ordered, dense atomic packing. By contrast, general high-angle random boundaries typically exhibit significantly larger free volumes and more severe atomic disorder.
Figure 5.
(a) The Voronoi volume of each point on the FeΣ3(111) grain boundary. (b,c) Fe and Nb Voronoi volume diagrams, respectively.
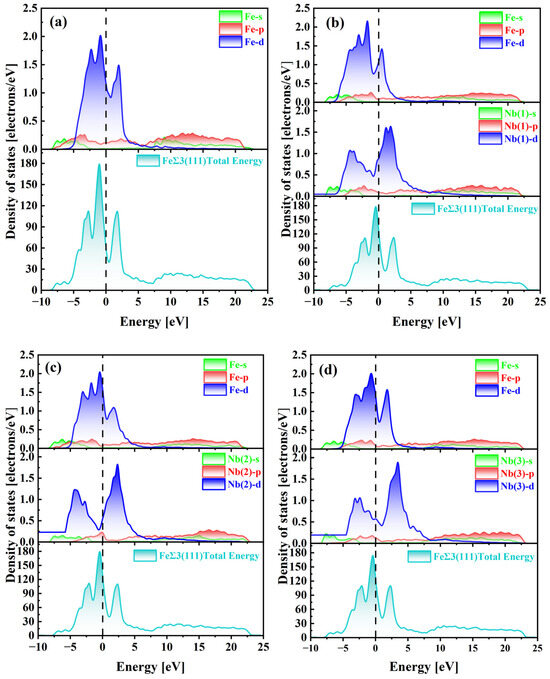
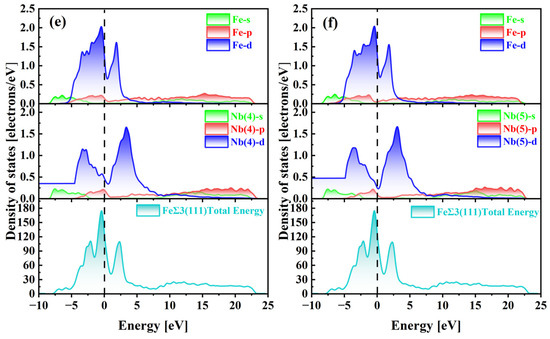
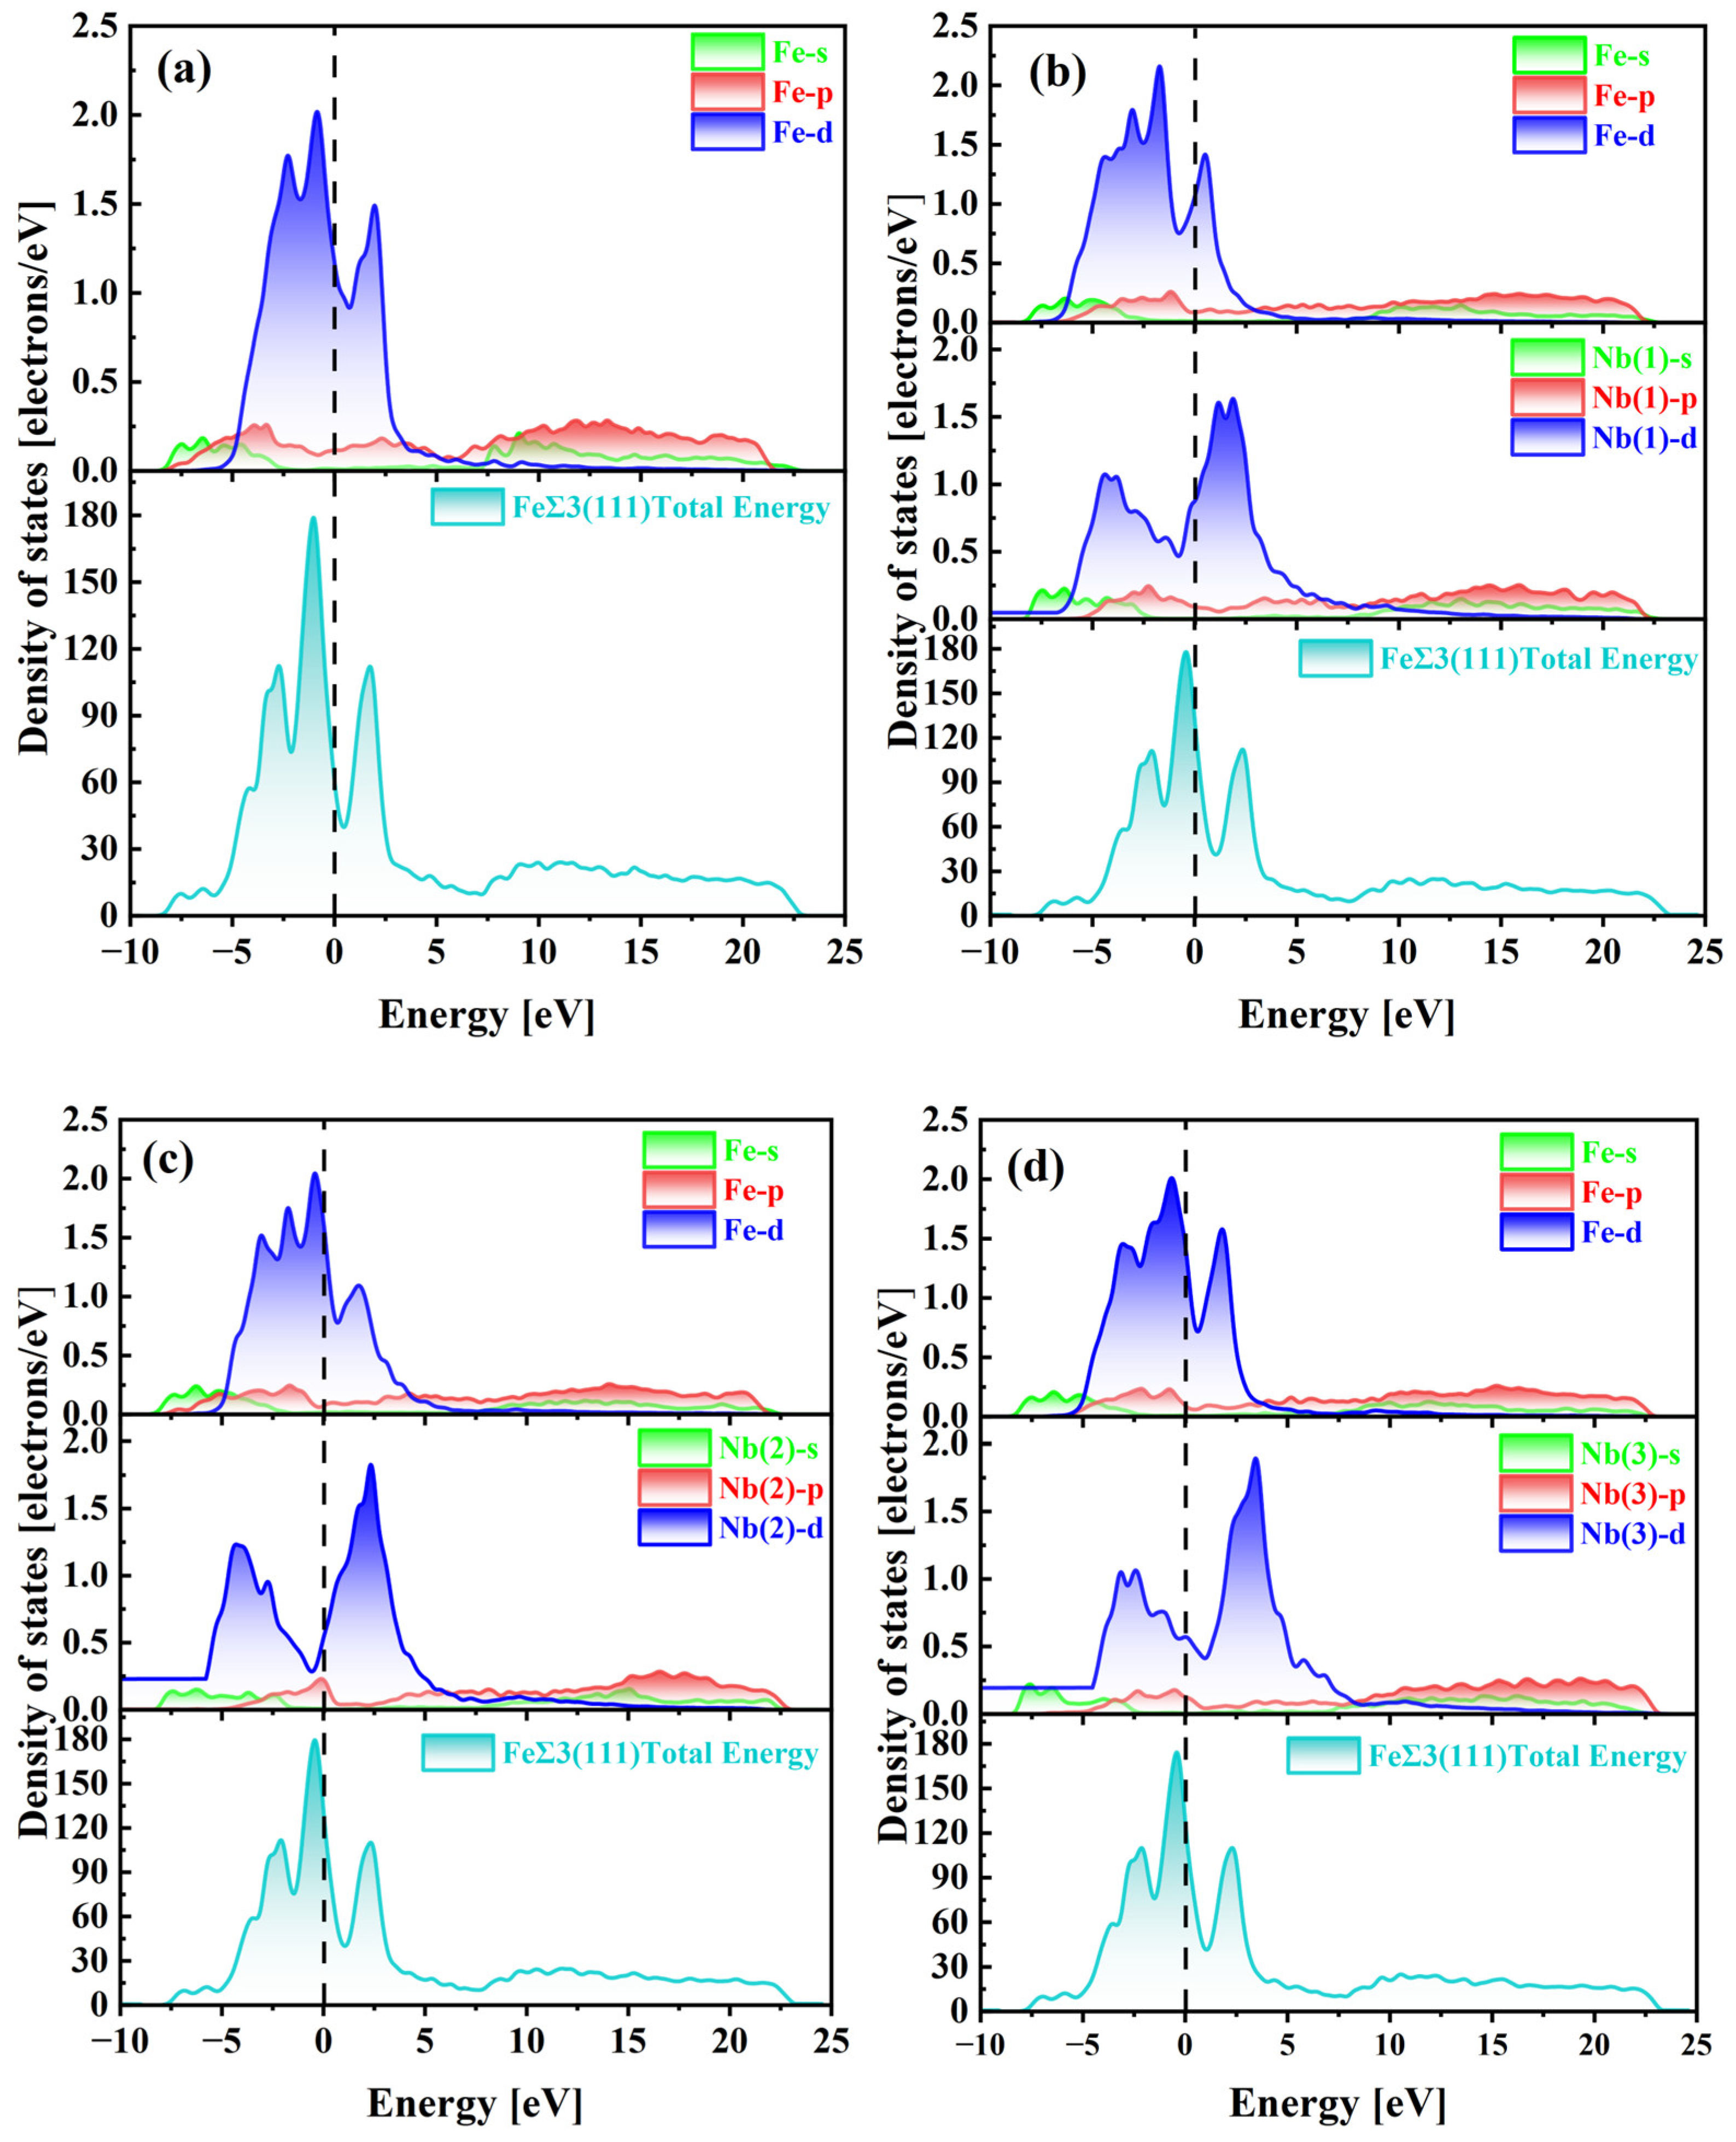
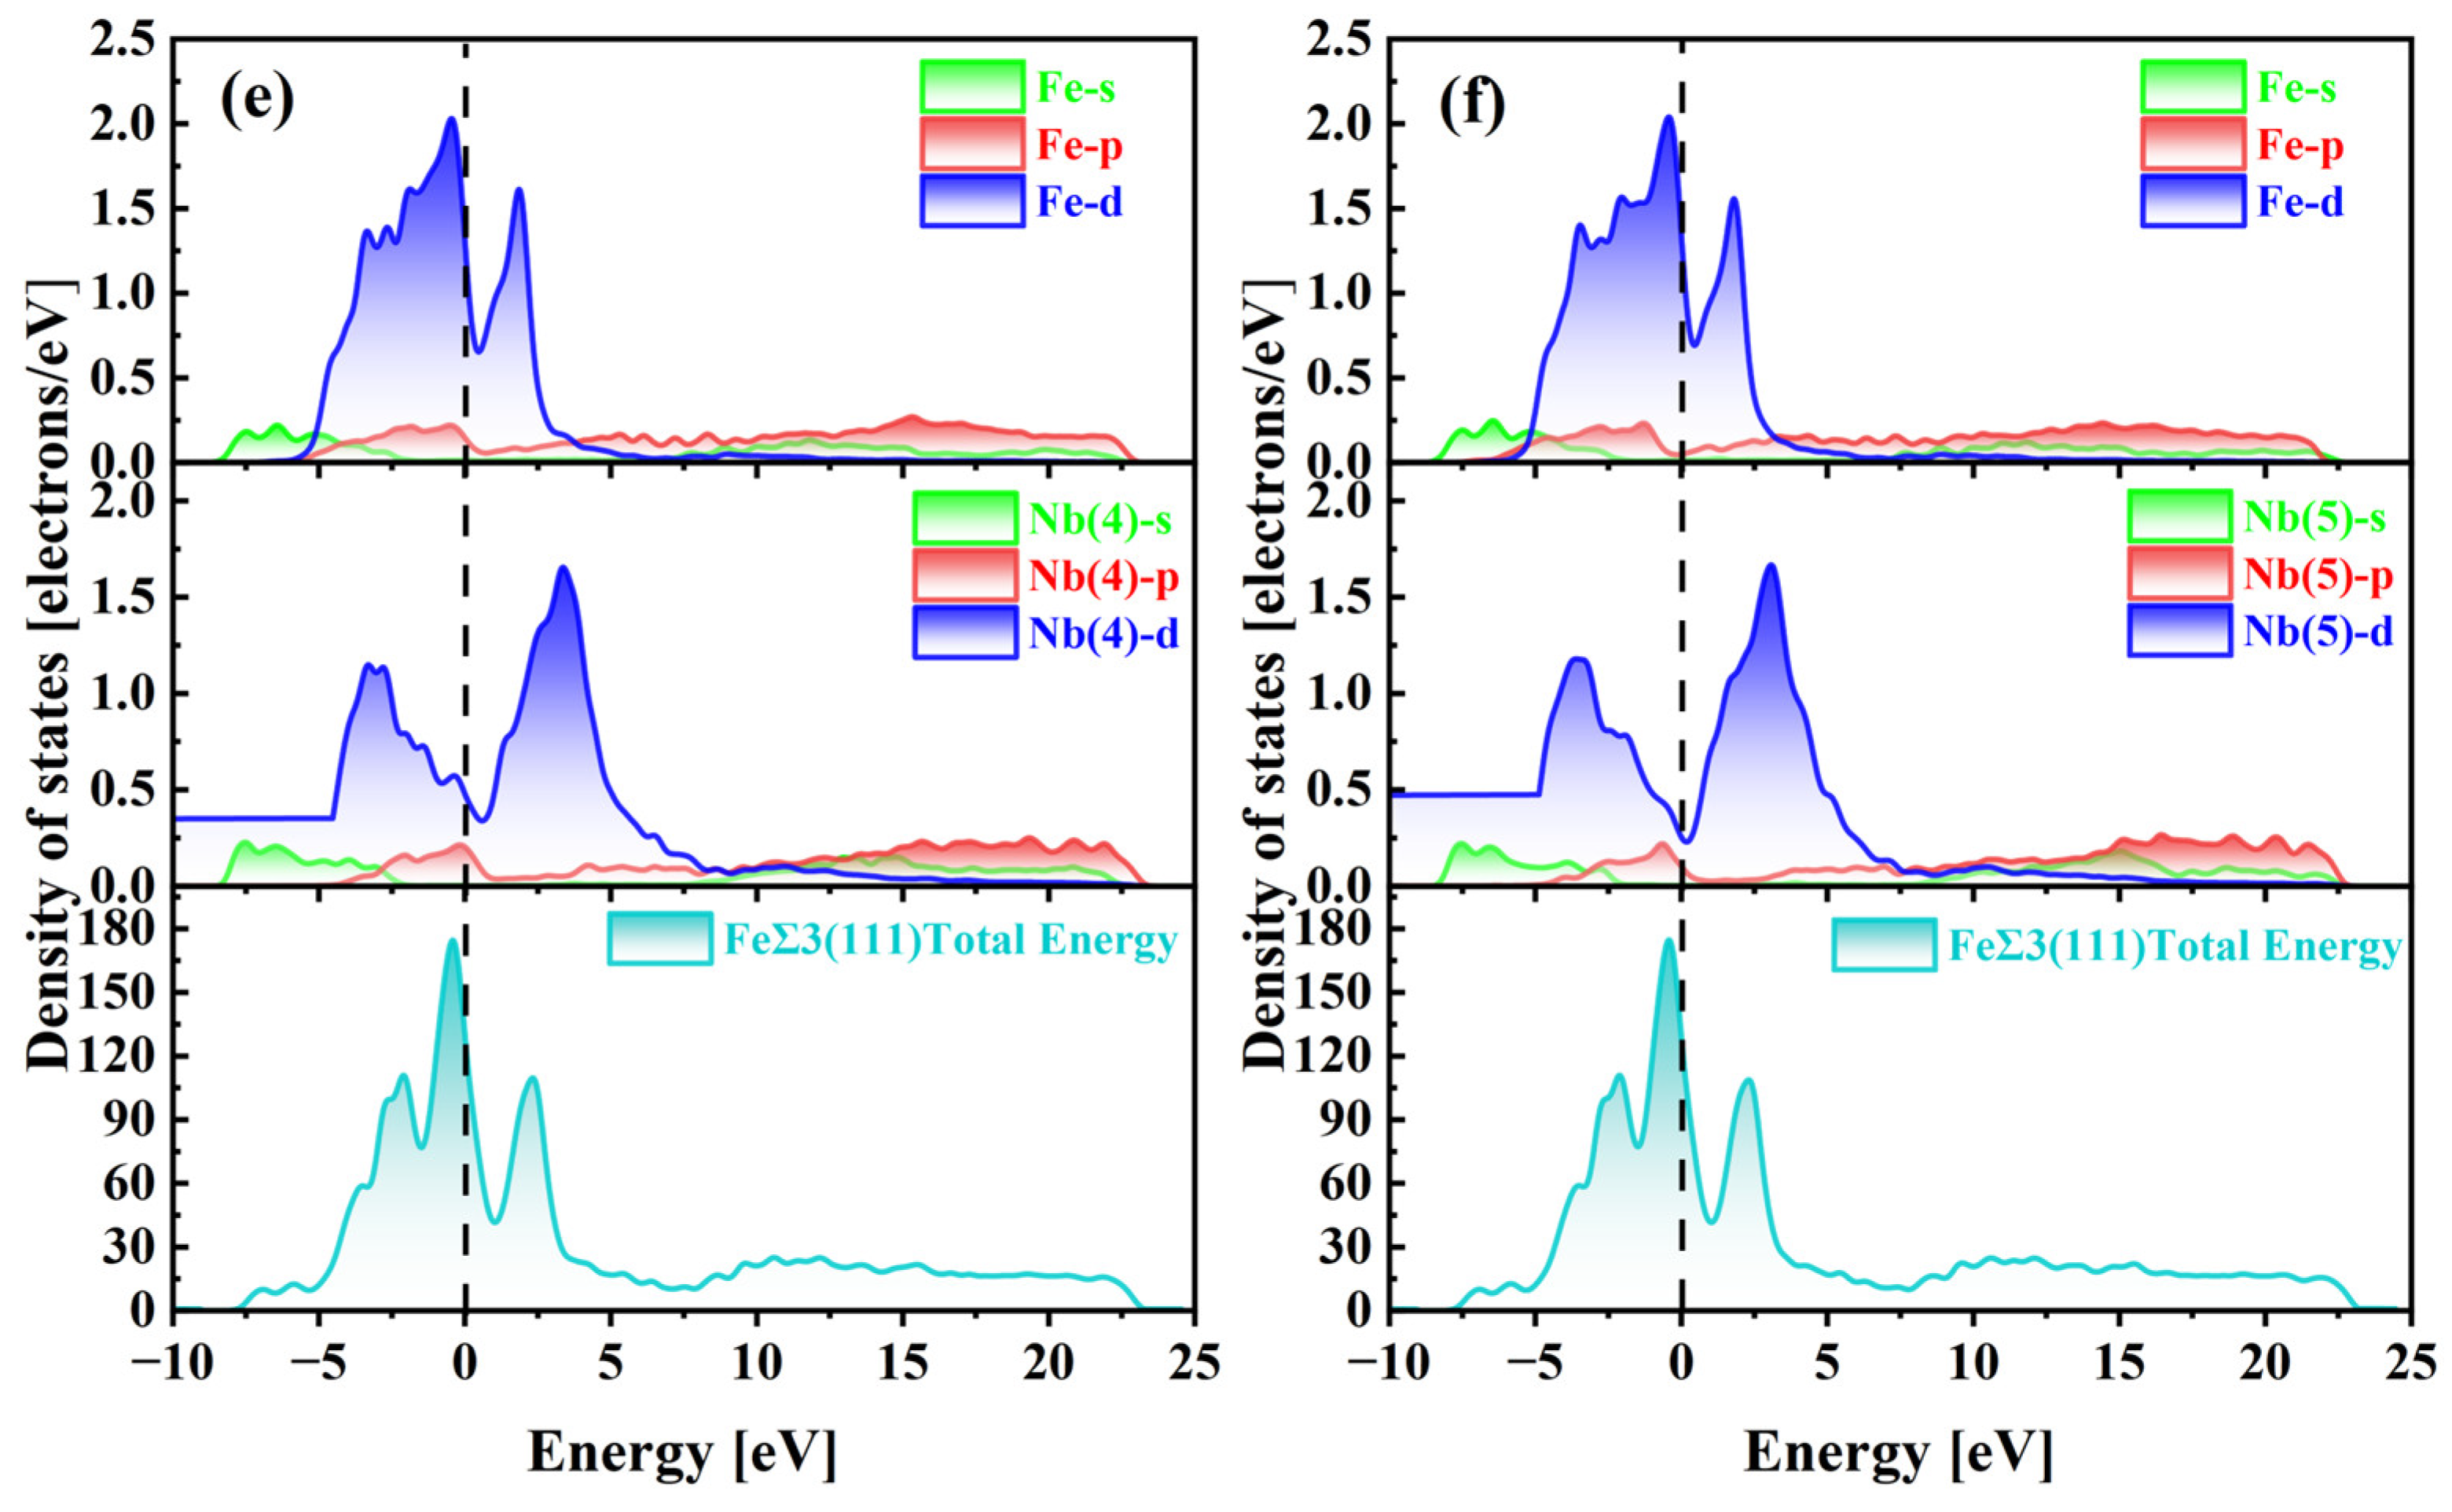
Figure 6 shows the density of states (DOS) of the pure FeΣ3(111)[1−10] grain boundary and of the grain boundary when Nb atoms were doped at different sites. Figure 6a represents the density of states at the grain boundary of pure FeΣ3(111), where the dashed line indicates the Fermi level. It can be seen that the density of states at the Fermi level was positive, confirming that this grain boundary structure exhibited metallic characteristics. The electronic states near the Fermi level mainly resulted from the contribution of the Fe-d orbitals. Figure 6b–f illustrates the density of states after the Nb atoms segregated at sites 1–5. Compared with Figure 6a, one of the most notable changes is that the density of states peaks corresponding to the d orbitals of Nb atoms appeared below the Fermi level, in the range from −2 eV to about −6 eV. Meanwhile, the density of states peaks from the Fe-d orbitals near the Fermi level generally decreased, which was most pronounced at site 1. In the energy range from −6 eV to 0 eV, the d-orbital density curves of Fe and Nb showed significant overlap and correspondence in shape, indicating that strong orbital hybridization occurred between the Fe-3d electrons and Nb-4d electrons in this energy range. Therefore, Nb atoms not only formed metallic bonds with neighboring Fe atoms but also developed directional covalent bonds, which enhanced the local bonding.
Figure 6.
The density of states in pure FeΣ3(111) grain boundary and Nb atoms at different sites. (a) The pure FeΣ3(111)[1−10] grain boundary. (b) Site 1. (c) Site 2. (d) Site 3. (e) Site 4. (f) Site 5.
The extent of electronic structure modification also differs among segregation sites. At site 1, the density of states of the Fe-d orbital at the Fermi level decreased the most, and the hybridization with the Nb-d orbital was most pronounced. From site 1 to site 5, this change effect gradually weakened. Furthermore, through comparison with Figure 6, it can be observed that the main changes were concentrated on the local density of states curves near the doping sites. This confirms the conclusion that the alloying element Nb directly interacted with the nearest Fe atoms in the vicinity through its own unique electronic properties, thereby influencing the local bonding characteristics.
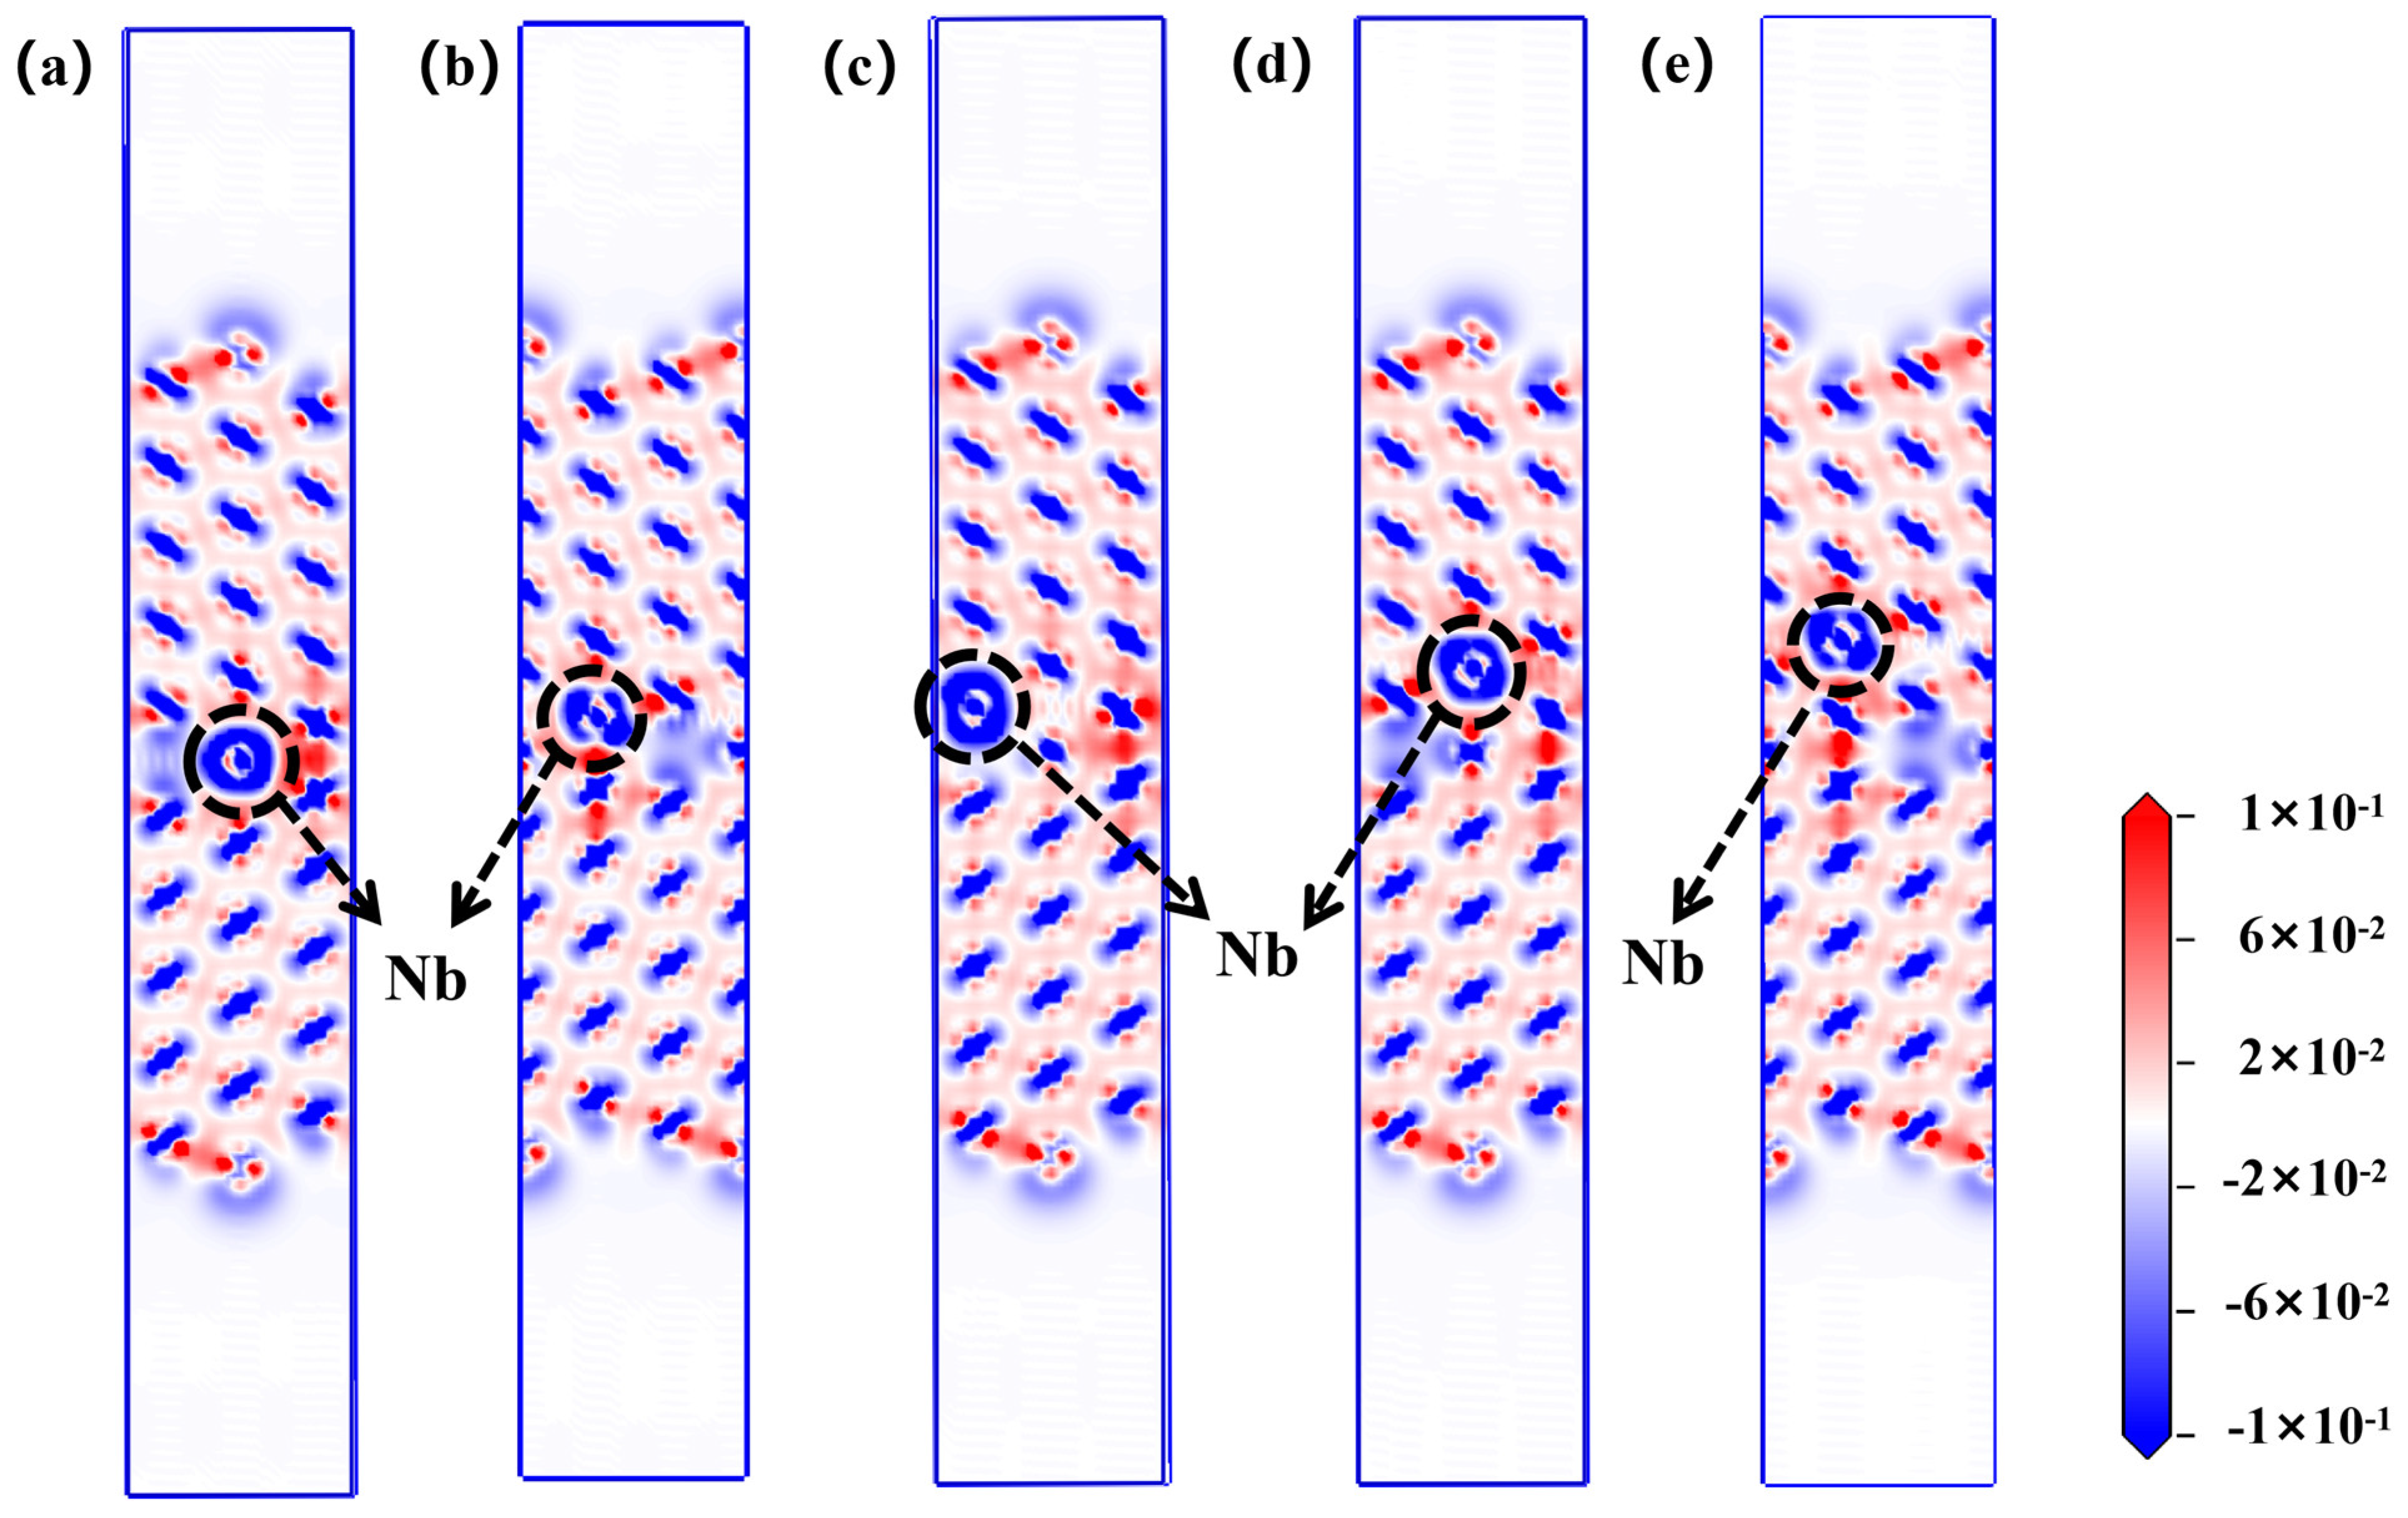
Figure 7 depicts the charge density difference following the doping of Nb atoms at different sites along the FeΣ3(111)[1−10] grain boundary. Among them, the red area indicates the aggregation of electrons, while the blue area represents the dissipation of electrons. Figure 7a–e clearly demonstrates that when Nb atoms were doped into five different sites at the grain boundaries, a distinct redistribution of charge occurred in their vicinity. Specifically, a red electron concentration zone appeared between the Nb atom and the neighboring Fe atom, connecting the two. Simultaneously, a blue electron depletion zone was typically observed near the nucleus of the Nb atom themselves. There were significant differences in the charge redistribution characteristics at different sites, which is highly consistent with previous results from energy and electronic structure calculations. At site 1, shown in Figure 7a, the electron cloud associated with the red electron cluster connecting the Nb and Fe atoms displayed the largest extent and most distinct shape, indicating that at site 1, the degree of electron sharing between Nb and the surrounding Fe atoms was highest, and the covalent bonding was strongest. The charge accumulation between Nb and Fe, while visible at all sites, exhibited the strongest localization and largest spatial extent at site 1, with a systematic attenuation toward site 5, consistent with the energetic and DOS trends.
Figure 7.
The charge density difference of Nb at different sites on the FeΣ3(111) grain boundary. (a) Site 1. (b) Site 2. (c) Site 3. (d) Site 4. (e) Site 5.
3.2. Grain Boundary-Strengthening Energy
After clarifying the segregation pattern of solute elements at grain boundaries, it is necessary to further investigate its impact on the mechanical properties. At present, the most widely used and highly accurate evaluation system for the influence of solute elements on the strength of grain boundaries is the grain boundary embrittlement model proposed by Rice and Wang [17]. The core of this model lies in the introduction of a key parameter known as embrittlement energy (). The positive or negative value of provides a qualitative prediction of whether solute segregation reduces or enhances the grain boundary cohesive strength. The calculation of the ∆ESE embrittlement factor is as follows:
In the formula, and refer to the total energy of the complete grain boundary supercell containing 62 atoms and 61 Fe + 1 Nb, comprising 15 atomic layers and vacuum slabs on both sides, respectively. and represent the total energy of the two resulting Fe(111) free surface supercells obtained by cleaving the grain boundary supercell along the interface plane (each containing half the atoms and vacuum slabs) with and without the segregated solute A, respectively. From Equation (3), it can be seen that the larger the absolute value of the negative value of ∆ESE, the more significant the enhancement effect of the segregated element on the bonding strength of the grain boundary. Conversely, a positive signifies that the segregation weakens the bonding ability of the grain boundary, leading to embrittlement.
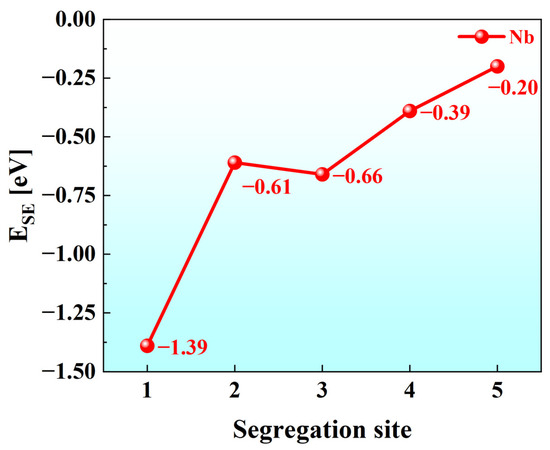
Figure 8 presents the calculated embrittlement energies for Nb atoms segregated at different sites of the FeΣ3(111) grain boundary. The values at the five sites were all negative, indicating that the segregation of Nb atoms contributed to enhancing the cohesive strength of the grain boundaries and thus exerted a strengthening effect. However, the degree of enhancement varied significantly with the segregation site. From site 1 to site 5, the values of the embrittlement energy showed a general increasing trend, with site 1 exhibiting the most negative value, suggesting a systematic weakening of the toughening effect with an increasing site number. By combining the results of the embrittlement energy with the segregation energy, it can be observed that the site with the most negative segregation energy (site 1, −2.47 eV) also exhibited the most negative embrittlement energy (−1.39 eV). This indicates that the Nb atom was thermodynamically most inclined to concentrate at a certain site, and this was precisely the site where it could exert the strongest cohesive strengthening effect on the grain boundaries. This correlation holds significant practical significance, indicating that not only do the Nb atoms undergo spontaneous segregation at the FeΣ3(111) grain boundary, but this segregation behavior is essentially a beneficial strengthening mechanism. By preferentially occupying specific grain boundary sites, Nb atoms can most effectively enhance the grain boundary cohesion, which may help improve the material’s resistance to intergranular failure during service.
Figure 8.
Embrittlement energy of Nb atoms at different sites on the FeΣ3(111) grain boundary.
3.3. Strain and Grain Boundary Behavior Analysis
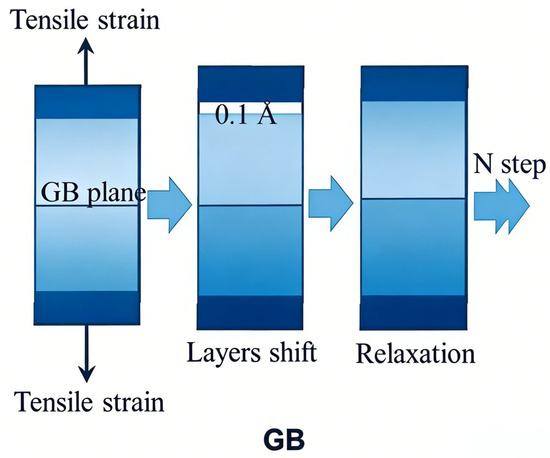
In this work, the simulation method for the first-principles tensile test adopted the optimized uniaxial deformation approach (referred to as OUD). As seen in Figure 9, this approach employs a layered crystal model, in which several atomic layers on the outer side of the grain boundary are fixed to preserve the properties of the bulk material. The lighter areas represent regions where atoms can relax, while the darker areas represent regions where atoms are fixed. After the unit cell is stretched incrementally to achieve a predetermined deformation, the coordinates of the remaining atoms are optimized to establish an energy–strain curve. When the unit cell dimensions perpendicular to the grain boundary approach the critical deformation threshold, spontaneous fracture occurs at the grain boundary, thereby allowing the critical strength of the grain boundary to be determined.
Figure 9.
Schematic diagram of the OUD tensile test scheme.
The optimized uniaxial deformation (OUD) tensile simulations were conducted by incrementally increasing the tensile strain along the [111] direction at a specified step size. At each strain step, the outermost four atomic layers on both sides of the grain boundary were fixed to preserve bulk-like boundary conditions, while all remaining internal atoms were fully relaxed. The Poisson effect was neglected by fixing the transverse lattice parameters (along [11−2] and [1−10]) to reduce the computational cost; this constraint is known to slightly overestimate the absolute stress values but preserves the qualitative fracture trend and the relative strengthening effect. The stress was derived from the CASTEP stress tensor output.
As shown above, Nb atoms preferentially segregated at site 1 on the FeΣ3(111) grain boundary and contributed to the strengthening of this grain boundary. Therefore, tensile experiment simulations would be conducted for the pure FeΣ3(111) grain boundary and the model structure with Nb segregated at site 1 of the grain boundary. In the experiment, a uniaxial tensile strain was applied and defined as follows:
Here, denotes the length of the supercell along the [111] direction when the tensile strain is , and denotes the initial length of the supercell along the [111] direction. In both cases, the vacuum layer thickness was excluded. To reduce the computational cost, the Poisson effect was neglected by fixing the transverse lattice parameters.
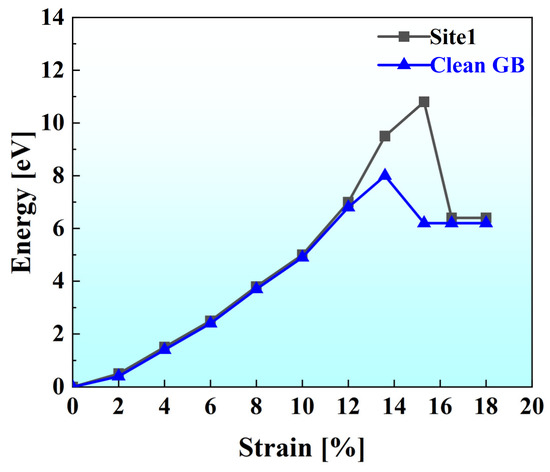
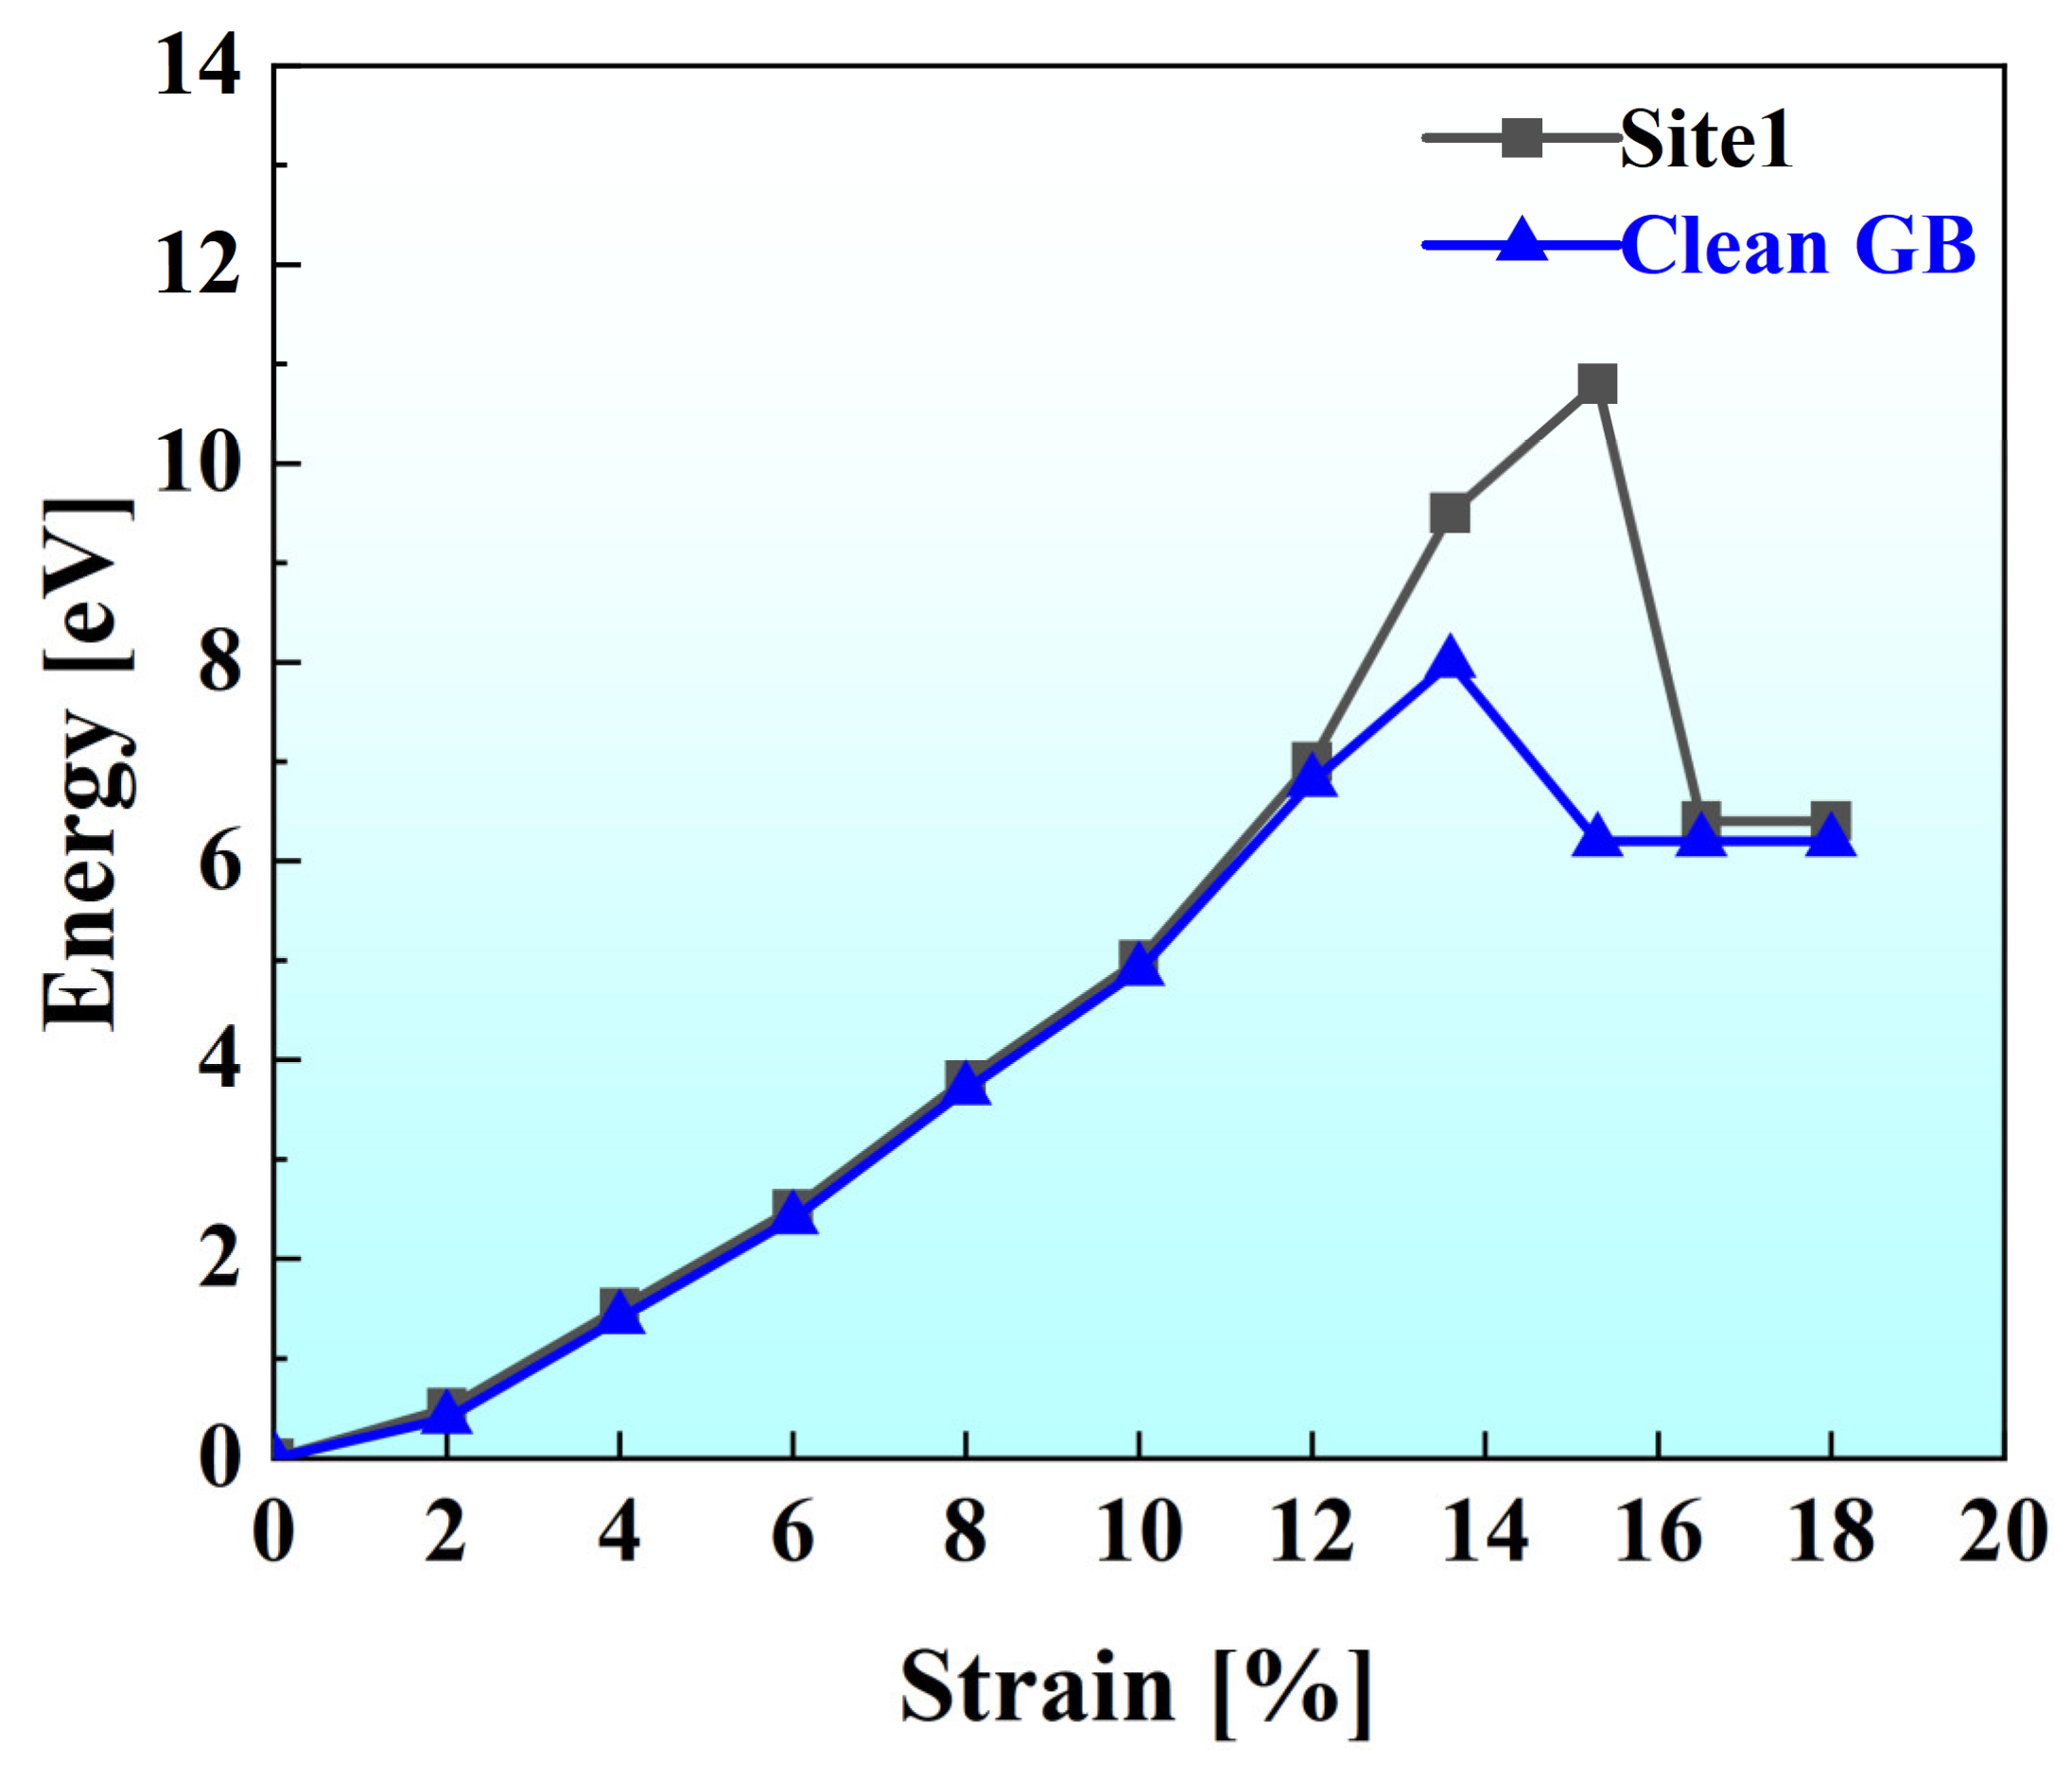
Figure 10 shows the energy–strain curves for two systems during tensile loading: one with a pure FeΣ3(111) grain boundary and the other with Nb atoms segregated at site 1 of that grain boundary. From the analysis of the variation patterns of the curves, both systems exhibited a characteristic two-stage variation in total energy with increasing tensile strain. In the case of the pristine FeΣ3(111) grain boundary system, the total energy steadily rose with increasing strain during the early tensile stage, due to increasing bond elongation and strain energy accumulation. When the tensile strain reached 13.58%, the total energy reached a maximum and then decreased sharply, indicating grain boundary decohesion, i.e., rupture of the grain boundary interface. Compared with the pure FeΣ3(111) grain boundary system, for the Nb-segregated grain boundary (Nb at site 1), the trend of total energy variation with the strain was different. Within the same range of stretching strain, the total energy of the heterogeneous system was higher overall than that of the pure system, and the strain at which the energy maximum occurred was 15.76%. This phenomenon indicates that the segregation of Nb atoms at site 1 could improve the energetic stability of the grain boundary, delay the occurrence of grain boundary rupture, and thereby play a strengthening role on the FeΣ3(111) grain boundary.
Figure 10.
Energy–strain curves of each system.
The theoretical tensile strength reported herein represents the minimum stress required to trigger mechanical instability in an ideal, defect-free, single-crystal grain boundary model at 0 K. It is an intrinsic materials property determined solely by the crystal configuration and interatomic bonding and therefore serves as an upper-bound estimate rather than a prediction of macroscopic yield strength. The absolute values (16–20 GPa) are orders of magnitude higher than experimentally accessible macroscopic strengths because the latter are governed by dislocation motion, microvoid nucleation, temperature, and strain rate effects that were not captured in the present ideal simulation. Nevertheless, the relative enhancement in theoretical strength and critical strain provides a physically meaningful atomistic metric for comparing grain boundary cohesion between the pristine and Nb-segregated systems. Therefore, it was necessary to obtain the stress–strain curve relationship during the simulation of the tensile process. The stress was obtained from Equation (5):
where is the stress exerted on the system, is the force acting on the system, and is the area of the grain boundary. The stress could also be derived from the energy variation with the strain as follows:
Combining Equations (5) and (6) yields
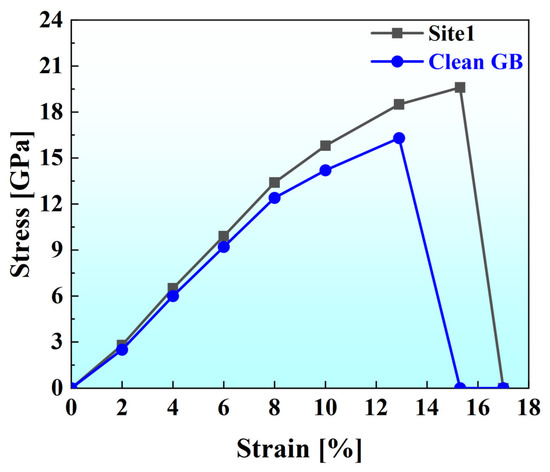
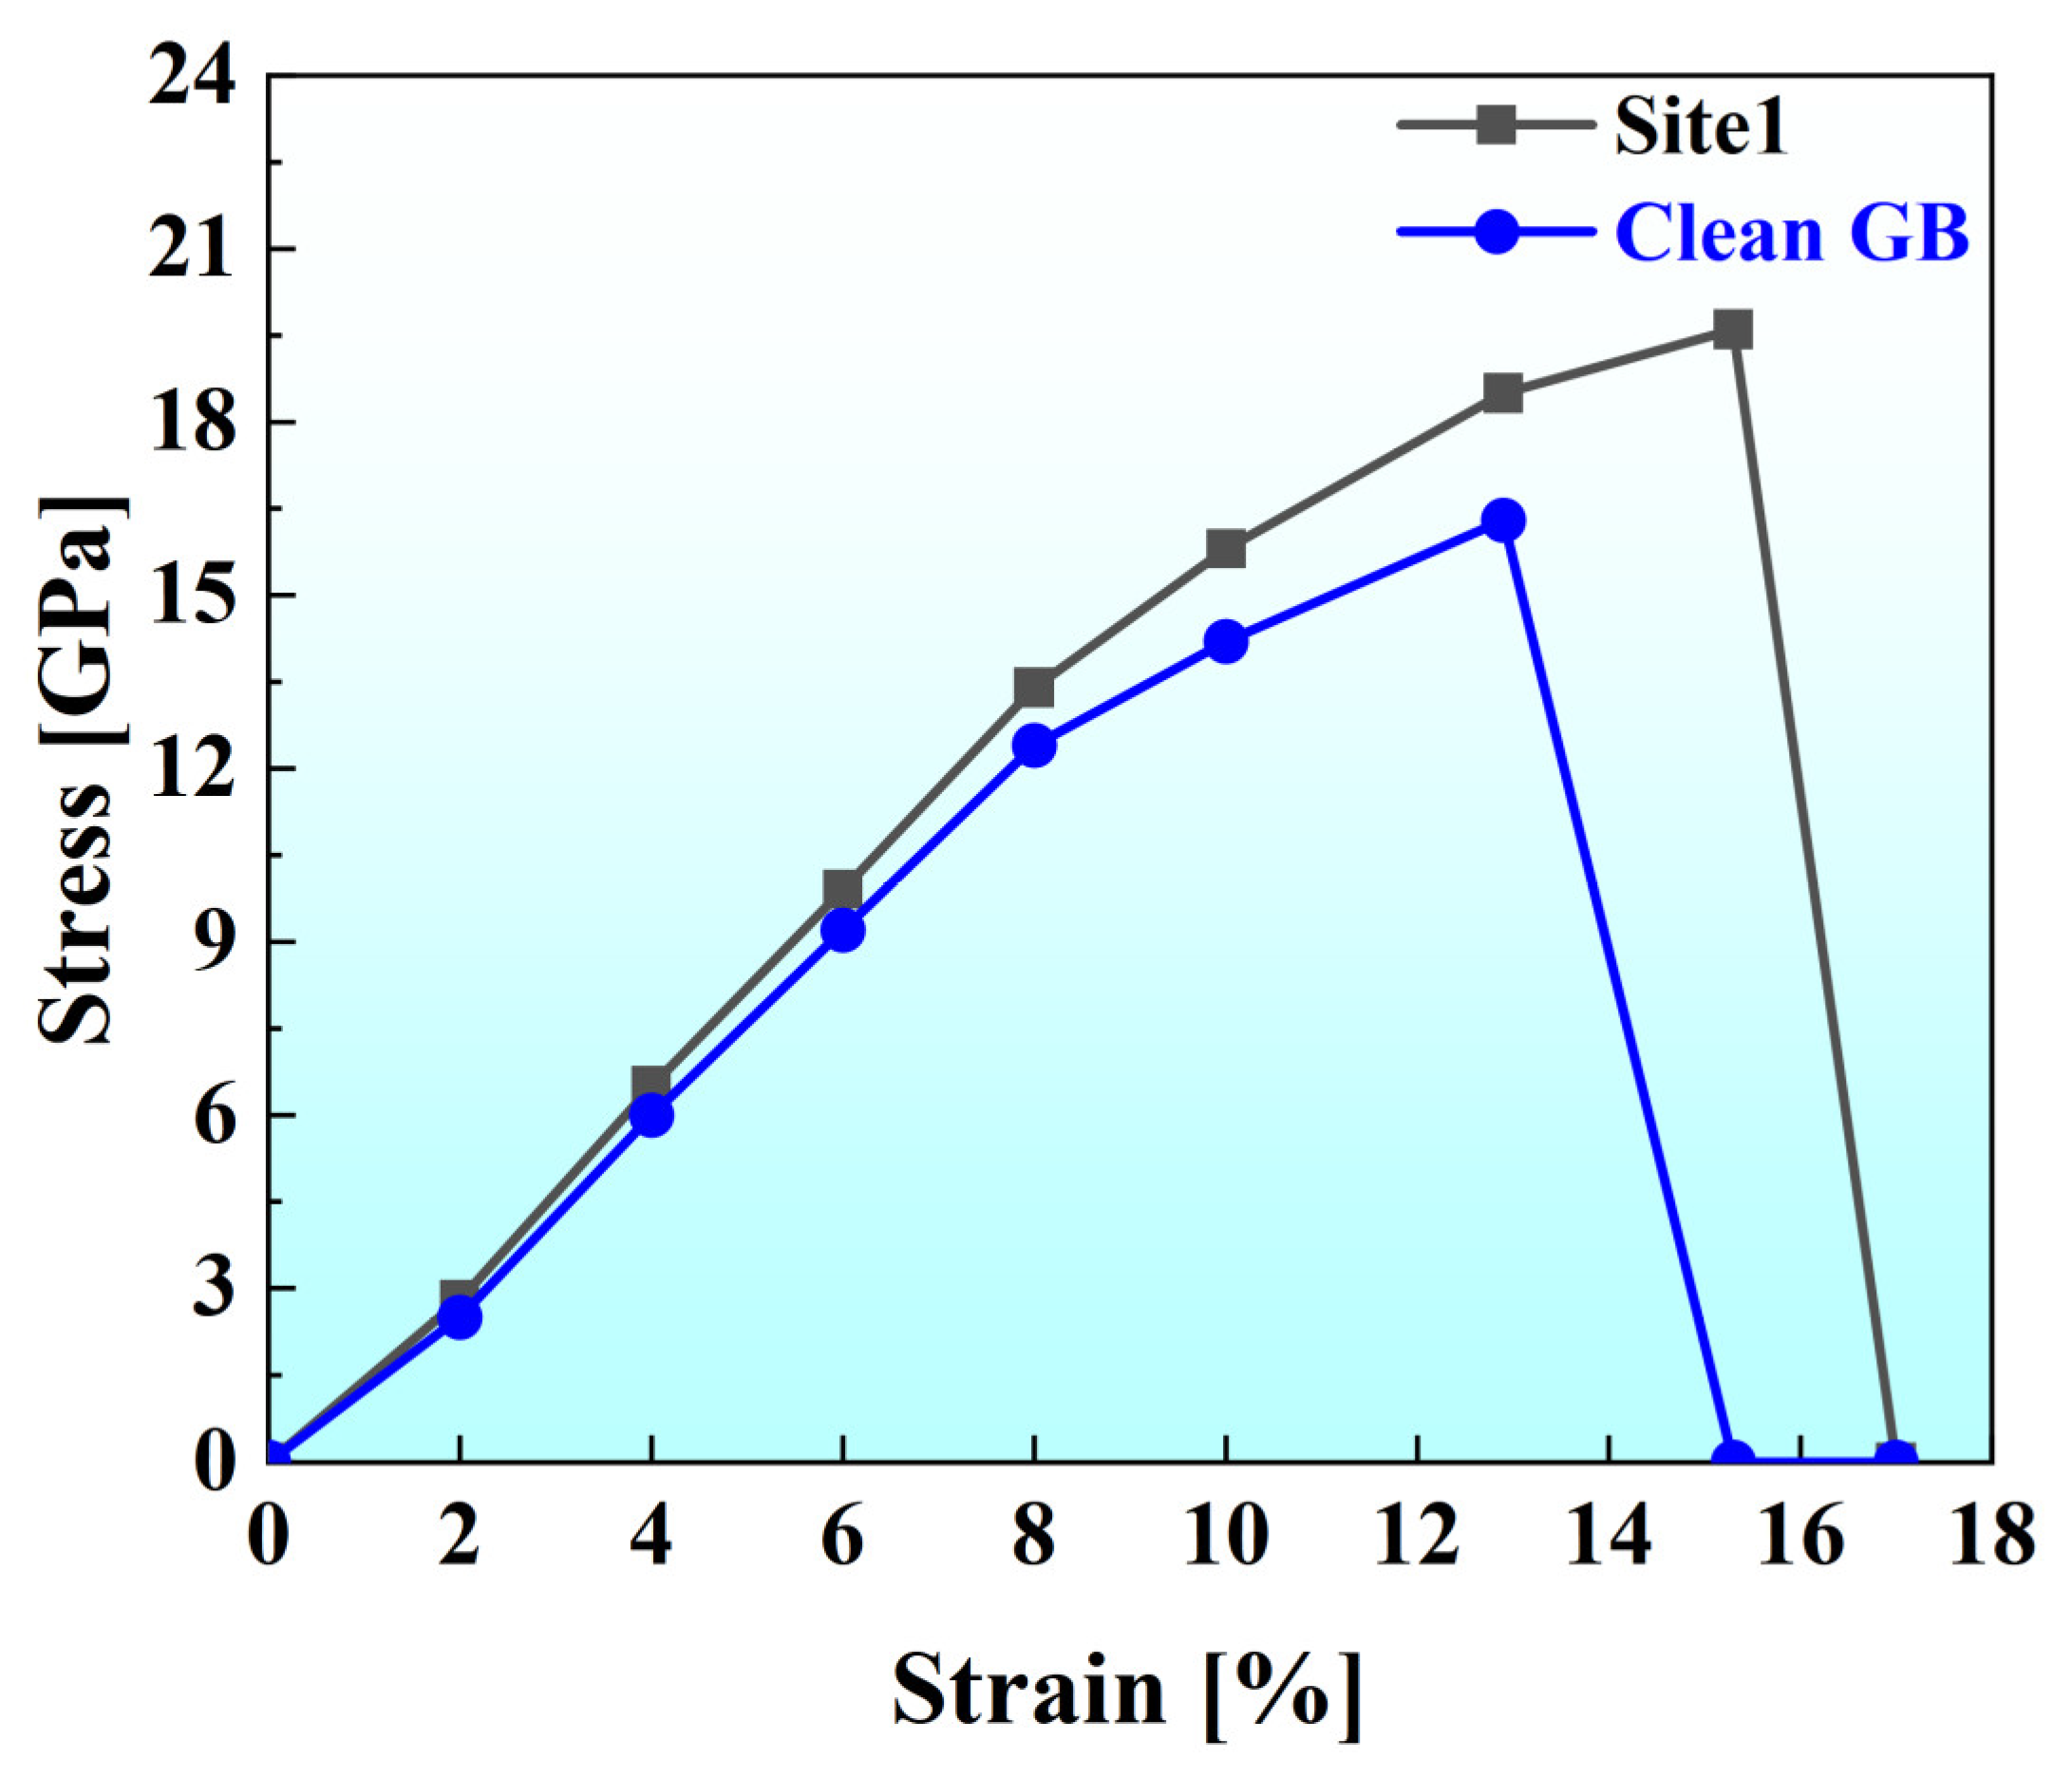
in which is the volume (Å3) of the system at strain . By incorporating Equation (7), the stress–strain response was directly obtained from the energy–strain curve. Depicted in Figure 11 is the comparative stress–strain response of the pristine FeΣ3(111) grain boundary and the Nb-segregated system (with Nb atoms occupying Site 1) subjected to tensile deformation. The two systems exhibited distinct behaviors. For the pristine FeΣ3(111) grain boundary system, at the pristine grain boundary, the stress increased monotonically with the strain during the early tensile stage due to progressive bond stretching, leading to stress accumulation. At a strain of 13.58%, the system attained its maximum stress, which corresponds to the theoretical tensile strength of the pristine FeΣ3(111) grain boundary, amounting to 16.32 GPa. Following the attainment of peak stress, the system stress sharply dropped to nearly zero, indicating grain boundary decohesion, consistent with the energy drop. This abrupt stress drop was consistent with the energy decrease shown in Figure 10; that is, when the strain exceeded 13.58%, the total energy of the system dropped rapidly, further confirming the grain boundary fracture. The system featuring Nb segregation at site 1 exhibited a stress–strain response that was uniformly superior to that of the pristine counterpart, with a peak stress of 19.64 GPa and a corresponding strain rising to 15.76%. This observation further corroborates Nb atom segregation at site 1 enhancing the resistance of the grain boundary to decohesion. The calculated enhancements in theoretical tensile strength (16.32~19.64 GPa) and critical strain (from 13.58% to 15.76%) provide an atomistic rationale for the improved ductility, correlating with the experimentally reported elongation of 22.3% at 900 °C. The segregation of Nb at the Σ3 twin boundary reinforced the intrinsic grain boundary cohesion, allowing greater deformation before fracture and thereby improving macroscopic ductility.
Figure 11.
The stress–strain curves of each system.
3.4. The Influence of Nb Segregation Concentration on the Strength of Grain Boundaries
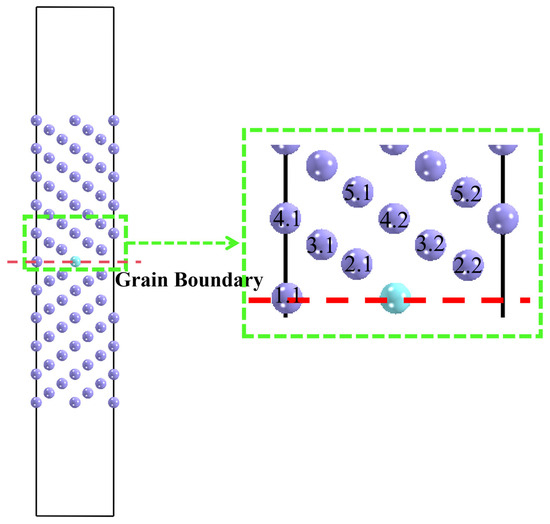
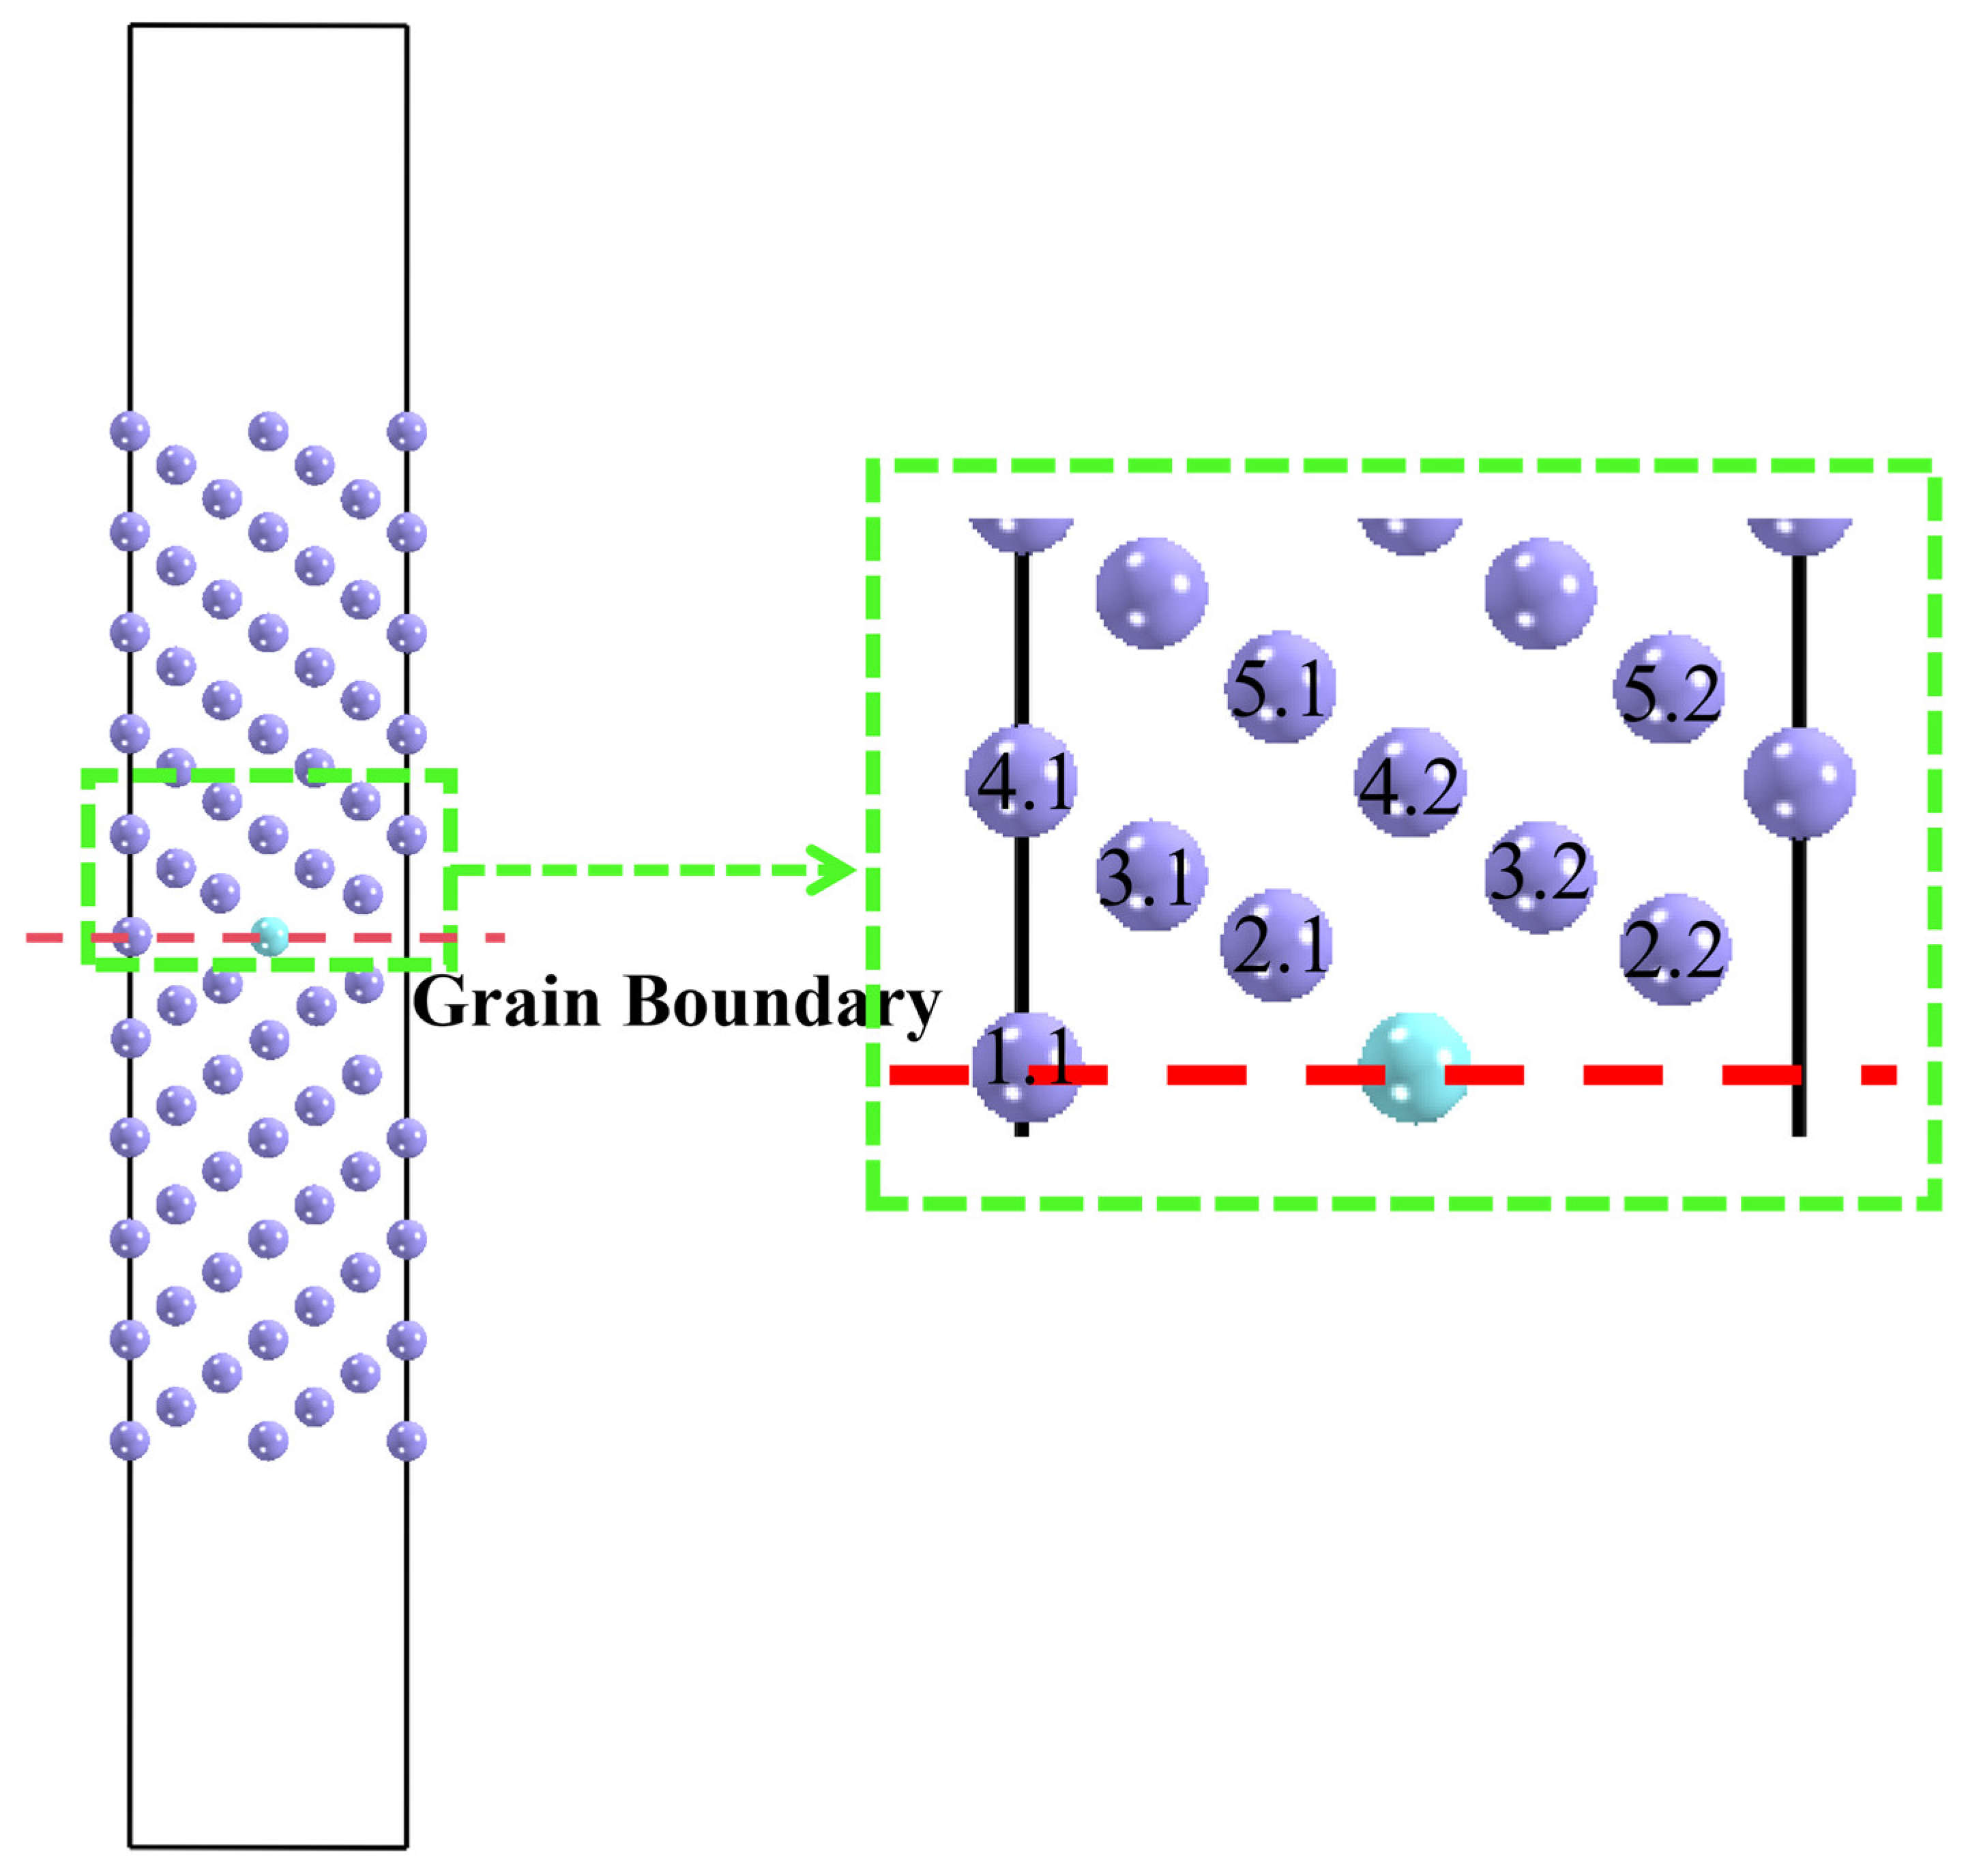
To further elucidate the mechanisms governing the grain boundary Nb segregation behavior, a quantitative analysis of Nb segregation was performed, aimed at delineating the relationship between Nb segregation and grain boundary strength, as well as its modulating effect on the boundary cohesion. As shown above, Nb exhibited a strong site preference at the Fe Σ3(111) grain boundary, and it preferentially occupied site 1. Accordingly, it became necessary to examine how the segregation of an additional Nb atom at different grain boundary positions perturbs the interfacial characteristics. Therefore, a total of nine different sites were selected for the study, as marked in Figure 12. The notation 1.1 denotes a configuration where the first Nb atom occupied site 1 and the second Nb atom was placed at an adjacent inequivalent site in the same grain boundary core region. All double-Nb configurations involved two distinct substitutional sites; no two Nb atoms shared the same lattice position.
Figure 12.
Nine segregation sites at the FeΣ3(111) grain boundary (the blue sphere represents an Nb atom).
As two Nb atoms segregate to the FeΣ3(111) grain boundary, the segregation energy cannot be directly derived from Equation (2). For the double Nb atom segregation system, the specific calculation formula for the segregation energy Eseg is as follows:
where is the total energy (eV) of the system featuring two Nb atoms segregated at the Fe Σ3(111) grain boundary, is the total energy of the clean FeΣ3(111) grain boundary, is the total energy of the system wherein one Nb atom is segregated into the Fe bulk lattice, and corresponds to the total energy of the pure Fe bulk.
The fracture energy is usually used as an indicator to evaluate the strength of grain boundaries. It is defined as the work required to break the grain boundary model along the crystal interface into two blocks with different orientations, followed by pulling them to infinity to form two free surfaces. The formula for calculating the fracture work is as follows [18]:
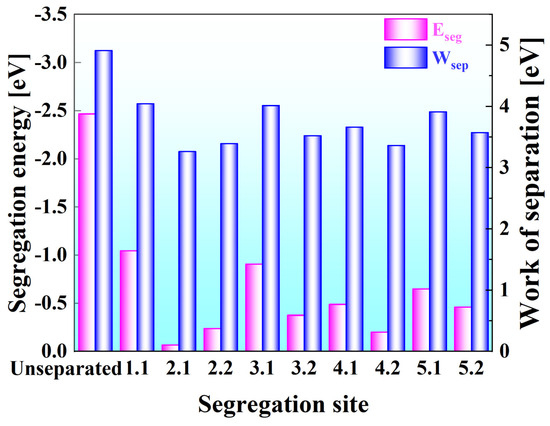
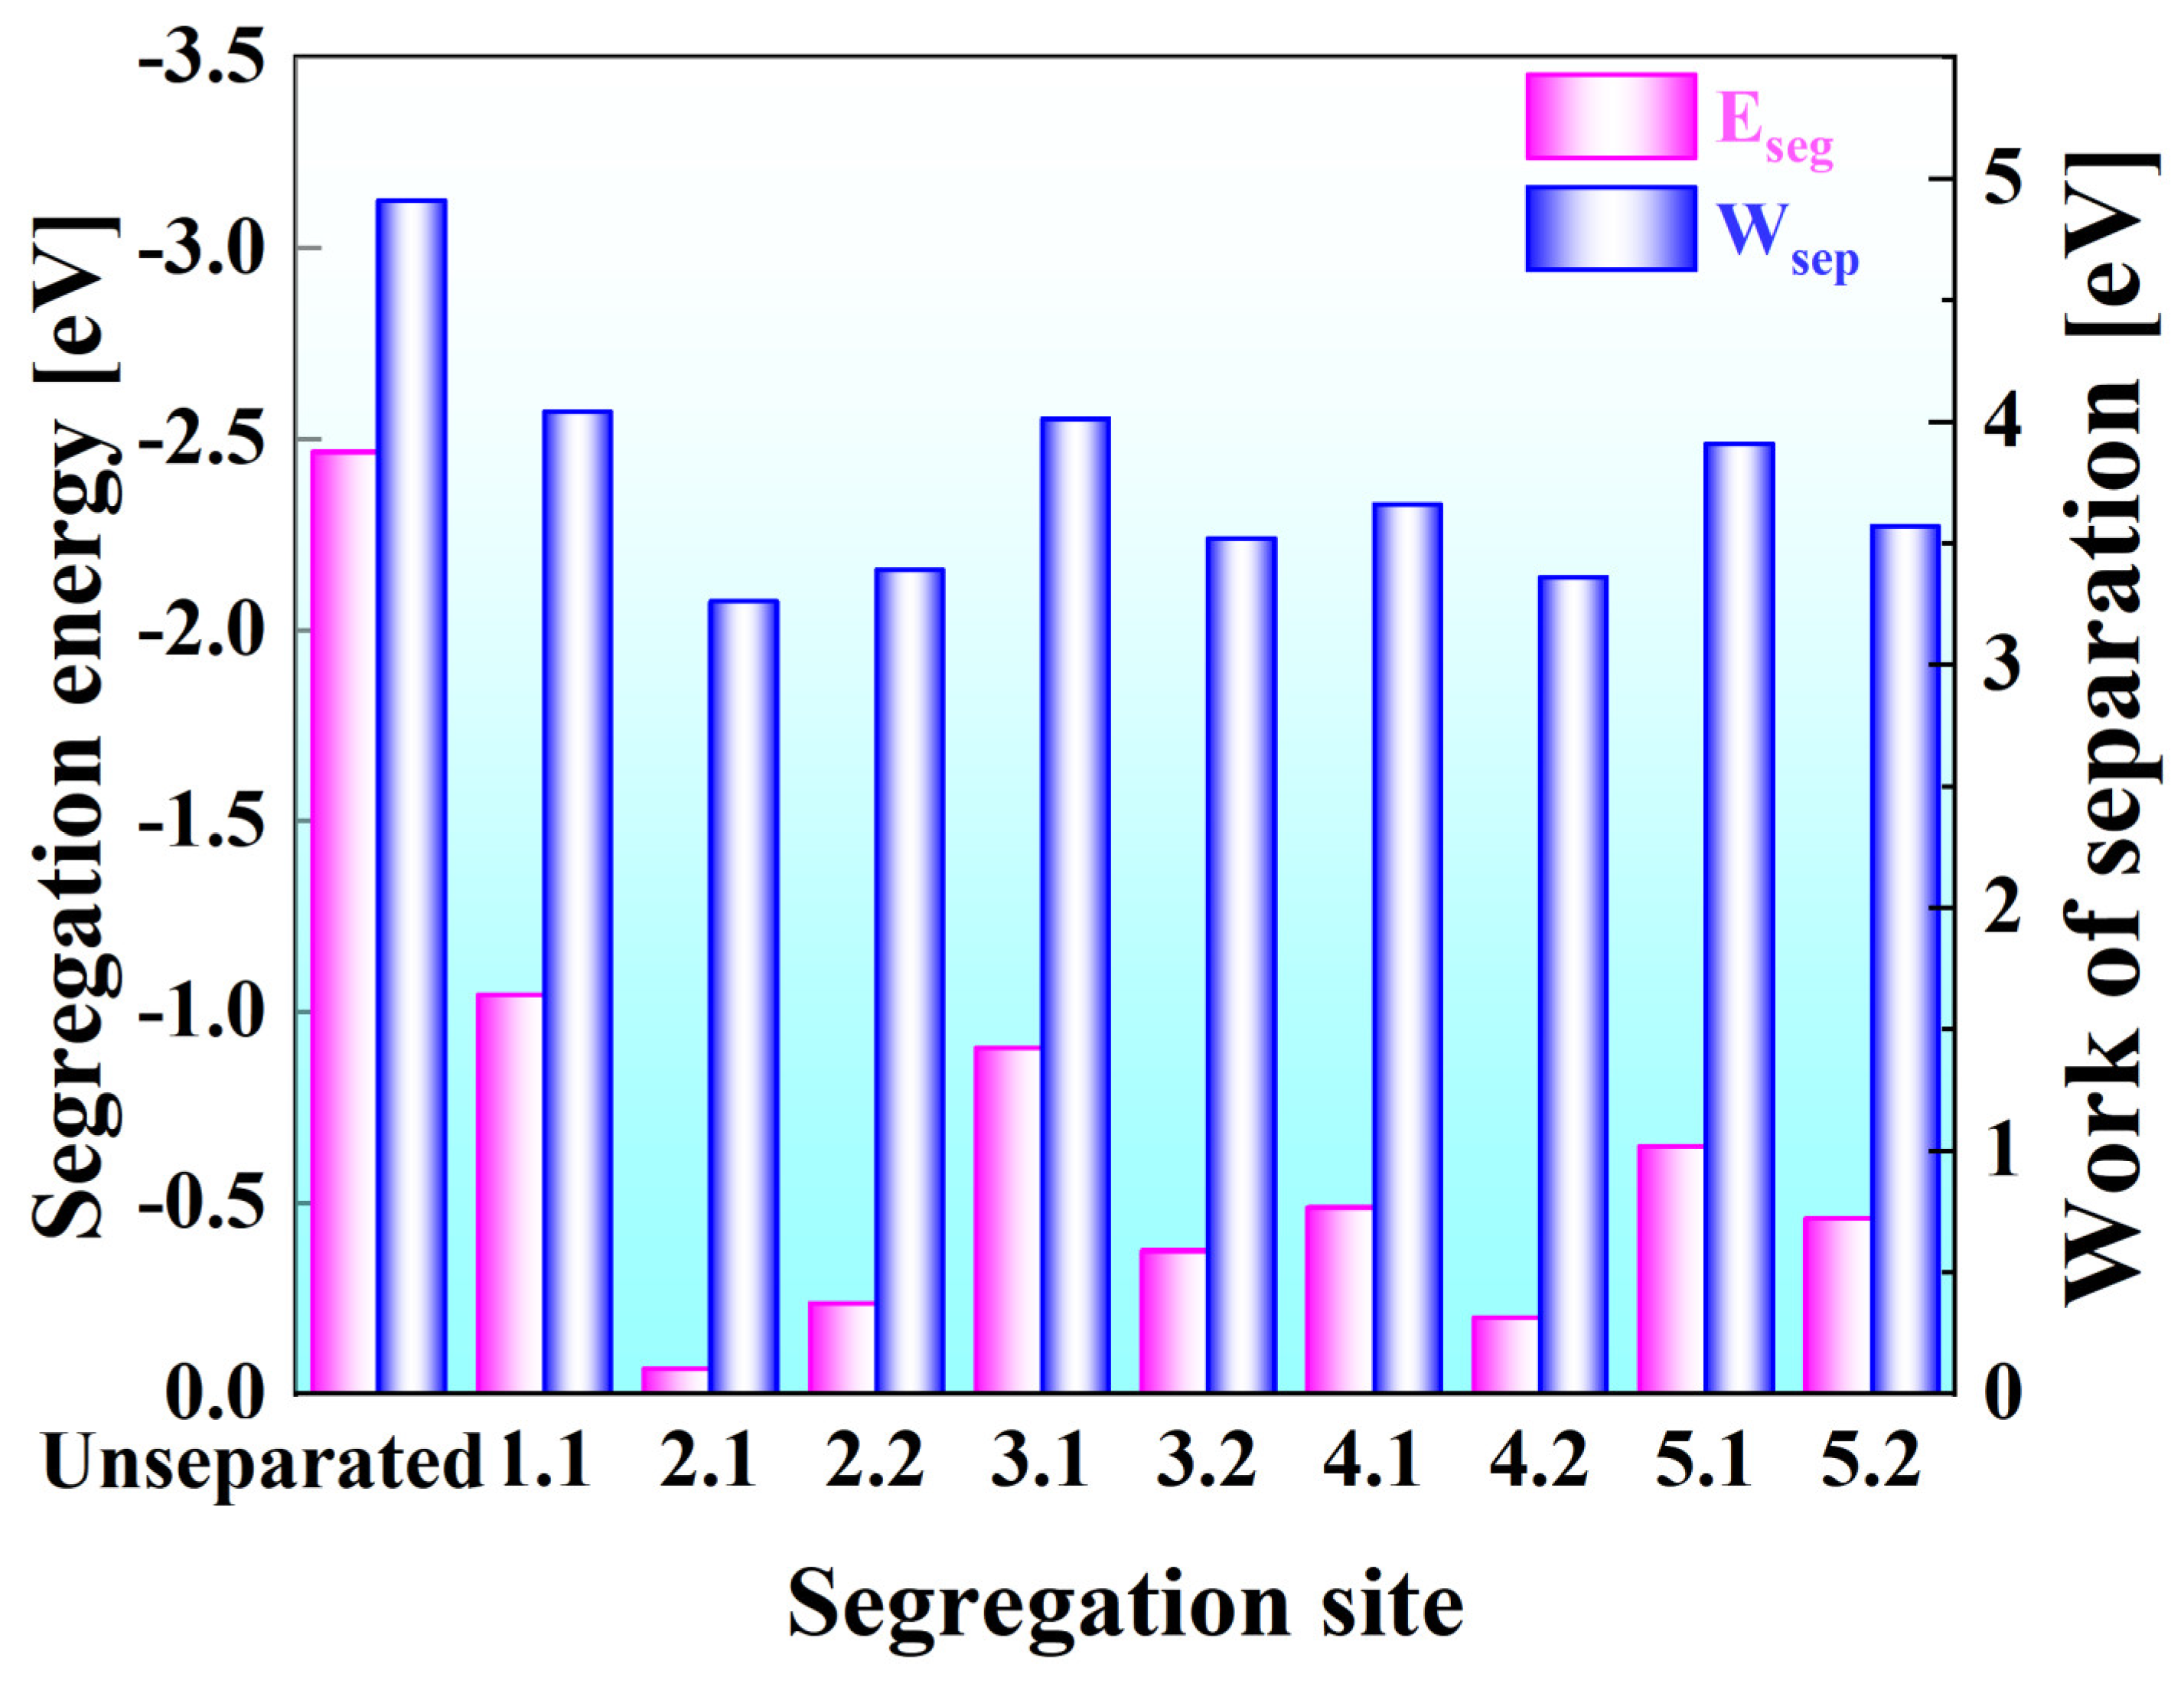
Here, and are the energies of the two free surfaces generated during grain boundary fracture, is the total energy of the FeΣ3(111) grain boundary, and S is the area of the grain interface. The calculated results of the segregation energy and fracture work of the second Nb atom at the nine sites on the grain boundary are listed in Table 2. The segregation energy of a single Nb atom at site 1 was −2.47 eV, being more negative compared with the diatomic segregation energies ranging from −0.07 to −1.24 eV. This indicates a pronounced tendency for the first Nb atom to segregate at site 1 of the grain boundary, consistent with the earlier conclusion regarding the strong segregation tendency of Nb at the FeΣ3(111) grain boundary.
Table 2.
The adsorption energy and strengthening energy of systems with two Nb atoms at different positions.
As shown in Figure 13, the segregation energies for the second Nb atom were significantly less negative, ranging from −0.07 to −1.24 eV, indicating that the driving force for the segregation of the second Nb atom was considerably weaker than that of the first. A clear site dependence of the segregation energy was observed. Site 1.1 displayed the most negative segregation energy (−1.24 eV), signifying the greatest driving force for segregation, while Site 2.1 yielded the least negative value (−0.07 eV), reflecting the most suppressed segregation propensity. The order of the partitioning energy tendency from strong to weak by site was 1.1 > 3.1 > 5.1 > 4.1 > 3.2 > 5.2 > 2.2 > 4.2 > 2.1.
Figure 13.
Segregation energies and works of fracture for two Nb atoms segregated at different sites.
The fracture energy of the diatomic segregation system ranged from 3.26 to 4.24 J/m2, which was lower than that for single-atom segregation. This indicates that the introduction of the second Nb atom generally reduced the strength of the grain boundary. Site 1.1, with a strong segregation tendency, enabled the Nb atoms to more effectively anchor in the core area of the grain boundary, partially offsetting the lattice distortion and bond weakening caused by the introduction of large solute atoms. Consequently, the reduction in grain boundary strength was relatively small. In contrast, for sites with a weak segregation tendency, such as site 2.1, Nb occupied a less favorable position and introduced a larger local stress field, which significantly weakened the grain boundary bonding and led to a substantial reduction in the fracture energy. The Nb atoms exhibited a competition between segregation-induced strengthening and solute-induced weakening at the FeΣ3(111) grain boundary. While the segregation of a single Nb atom unambiguously strengthens the FeΣ3(111) grain boundary, the present study reveals that the introduction of a second Nb atom generally reverses this trend, reducing the work of separation. This behavior indicates a competitive interplay between segregation-induced strengthening and solute-induced weakening. At low segregation levels, the electronic hybridization between Nb and Fe dominated, enhancing cohesion. At higher local concentrations, the cumulative elastic strain field around the oversized Nb atoms and possible Nb–Nb repulsive interactions became dominant, degrading the interfacial bonding. It is acknowledged that real grain boundaries in engineering materials may accommodate multiple segregated atoms or even continuous solute layers. The two-atom model employed here represents a tractable first step toward understanding this competitive behavior, capturing the transition from isolated solute strengthening to concentrated solute weakening. The results imply the existence of an optimal Nb concentration window at the grain boundary, beyond which further accumulation becomes detrimental.
The present study is subject to several limitations that should be acknowledged. First, all calculations were performed at 0 K, neglecting finite-temperature entropic contributions, magnetic fluctuations, and phonon effects. Second, the investigation was restricted to a single special low-energy grain boundary; generalization to random high-angle boundaries requires future comparative studies on Σ5, Σ7, and non-coincident site-lattice boundaries. Third, the possible competition between Nb segregation and Nb(C,N) precipitation was not considered. Fourth and finally, the OUD tensile simulation neglected the Poisson effect, dislocation activity, and plastic relaxation mechanisms that dominate macroscopic deformation.
Future work should address these limitations by employing finite-temperature ab initio molecular dynamics or Metropolis Monte Carlo methods to assess configurational entropy, extending the analysis to multiple grain-boundary types, and investigating co-segregation of Nb with C and N atoms. From an engineering perspective, the results suggest that strict control of the Nb content in 442D ferritic stainless steel (within the 0.3–0.6 wt.% range) and optimization of the annealing parameters to favor single-atom segregation at Σ3 twin boundaries may maximize grain boundary cohesion and ductility.
4. Conclusions
Based on the first-principles calculations, the segregation behavior of Nb at the FeΣ3(111) grain boundary and its effects on the grain boundary strength were investigated. The main conclusions are as follows:
- (1)
- The Nb atoms exhibited a strong site preference at the FeΣ3(111) grain boundary. The calculation results indicate that site 1 had the lowest segregation energy (−2.47 eV) and was the most thermodynamically stable favorable site, which correlated with its larger Voronoi volume and more open local atomic environment. This was consistent with the grain boundary angles of 50–60° observed in the experiment at 900 °C, indicating that the Σ3 twinned boundaries formed during recrystallization provided ideal sites for Nb segregation.
- (2)
- The analyses of the density of states and charge density difference revealed the electronic layer variation of Nb segregation. The Nb-4d and Fe-3d orbitals underwent significant hybridization in the energy range from −2 eV to the Fermi level, leading to localized charge accumulation between Nb and Fe atoms and strengthening the covalent bonding of Nb-Fe, which strengthened the grain boundary.
- (3)
- The uniaxial tensile via first-principles tensile simulation shows that Nb segregation increased the critical strain of the FeΣ3(111) grain boundary from 13.58% to 15.76% and the theoretical tensile strength from 16.32 GPa to 19.64 GPa. The calculated segregation energies were negative for all sites, with site 1 exhibiting the strongest effect (−1.39 eV).
- (4)
- Research on double Nb atom dislocations indicates that the introduction of a second Nb atom generally reduced the grain boundary strength, with the work of separation decreasing from 4.91 J/m2 in the case of a single Nb atom to 3.26–4.24 J/m2. The sites with more negative segregation energy caused less of a reduction in grain boundary strength, indicating a competition between segregation-induced strengthening and solute-induced weakening.
Author Contributions
Conceptualization, X.T.; methodology, X.L. (Xuefeng Lu) and X.T.; software, J.S. (Jiawei Shen); validation, L.Z.; formal analysis, L.Z.; investigation, X.L. (Xiaoyang Luo) and J.S. (Jie Sheng); resources, X.L. (Xiaoyang Luo); data curation, J.S. (Jiawei Shen) and J.S. (Jie Sheng); writing—original draft preparation, L.Z.; writing—review and editing, X.L. (Xiaoyang Luo), J.S. (Jiawei Shen) and J.S. (Jie Sheng); visualization, X.T.; supervision, X.L. (Xuefeng Lu); project administration, X.T.; funding acquisition, X.L. (Xuefeng Lu). All authors have read and agreed to the published version of the manuscript.
Funding
This research was funded by Longyuan Youth Talent Project of Gansu Province, Science and Technology Project of Jiayuguan City (24-01, QKJ24-05).
Data Availability Statement
The original contributions presented in this study are included in the article. Further inquiries can be directed to the corresponding author.
Conflicts of Interest
Lei Zhang and Xiaoyang Luo were employed by the Jiuquan Iron and Steel (Group) Co., Ltd. The remaining authors declare that the research was conducted in the absence of any commercial or financial relationships that could be construed as a potential conflict of interest.
Abbreviations and Nomenclature
| APT | atom probe tomography |
| bcc | body-centered cubic |
| DFT | density functional theory |
| DOS | density of states |
| FS | free surface |
| GB | grain boundary |
| GGA | generalized gradient approximation |
| HRTEM | high-resolution transmission electron microscopy |
| OUD | optimized uniaxial deformation |
| PBE | Perdew–Burke–Ernzerhof |
| pDOS | projected density of states |
| SCF | self-consistent field |
| Symbols | |
| A | grain boundary cross-sectional area [m2] |
| ΔESE | embrittlement [segregation-induced strengthening] energy [eV] |
| Ebulk | total energy of pure Fe bulk supercell [eV] |
| total energy of Fe bulk supercell with solute A [eV] | |
| EGB | total energy of clean grain boundary supercell [eV] |
| total energy of grain boundary supercell with solute A [eV] | |
| EFS | total energy of clean free surface supercell [eV] |
| total energy of free surface supercell with solute A [eV] | |
| Eseg | segregation energy [eV] |
| EF | Fermi energy [eV] |
| ε | tensile strain [%] |
| γGB | grain boundary energy [J/m2] |
| initial supercell length along [111] excluding vacuum [Å] | |
| supercell length at strain ε along [111] excluding vacuum [Å] | |
| NFe | number of Fe atoms in the grain boundary model |
| σ | theoretical tensile stress [GPa] |
| S | grain boundary interface area [m2] |
| system volume at strain ε [Å3] | |
| Wsep | work of separation or fracture energy [J/m2] |
References
- Hu, S.; Han, E.H.; Liu, X.B. Atomic-scale evidence for the intergranular corrosion mechanism induced by co-segregation of low-chromium ferritic stainless steel. Corros. Sci. 2021, 189, 10958. [Google Scholar] [CrossRef] [Scilit]
- Ma, X.; Zhao, J.; Du, W.; Zhang, X.; Jiang, L.; Jiang, Z. Quantification of texture-induced ridging in ferritic stainless steels 430 and 430LR during tensile deformation. J. Mater. Res. Technol. 2019, 8, 2041–2051. [Google Scholar] [CrossRef] [Scilit]
- Yan, H.T.; Bi, H.Y.; Li, X.; Zhou, X. Influence of Nb on Microstructure and Mechanical Properties of 0Cr11 Ferritic Stainless Steel. Iron Steel 2009, 44, 59–62+94. [Google Scholar]
- Xia, D.X. Application of Nb in the modern ferritic stainless steel. Shandong Sci. 2019, 32, 57–67. [Google Scholar] [CrossRef]
- Jang, J.H.; Kim, S.D. Evolution from clusters to precipitates in niobium-micro-alloyed ferritic steel: A combined in situ scanning transmission electron microscopy and atomistic study. Scr. Mater. 2024, 243, 115967. [Google Scholar] [CrossRef] [Scilit]
- Yan, X.Y.; Wang, H.B.; Zhang, C.D.; Wu, S.W.; Wang, Y.Q.; Cao, G.M.; Liu, Z.Y. Development of machine learning models for precipitation and recrystallization behavior in microalloyed steels. Iron Steel 2009, 60, 215–224. [Google Scholar] [CrossRef]
- Mai, H.L.; Cui, X.-Y.; Scheiber, D.; Romaner, L.; Ringer, S.P. The segregation of transition metals to iron grain boundaries and their effects on cohesion. Acta Mater. 2022, 231, 117902. [Google Scholar] [CrossRef] [Scilit]
- Li, Y.; Chen, W.; Li, Y.K.; Chen, Y.; Zhang, Q.-J.; Sun, L.-G.; Xiao, P.-C. Revealing the interface energy and electronic properties of NbN/α-Fe in shipbuilding steels. Mater. Today Commun. 2023, 35, 105852. [Google Scholar] [CrossRef] [Scilit]
- Li, Y.F.; Wang, C.B.; Hui, Y.; Chen, C.; Lu, X.; Sheng, J.; Tang, X. The influence of annealing on microstructure evolution and mechanical properties of 442 ferritic stainless steel. Metals 2026, 16, 167. [Google Scholar] [CrossRef] [Scilit]
- Bhattacharya, S.K.; Tanaka, S.; Shiihara, Y.; Kohyama, M. Ab initio perspective of the <110> symmetrical tilt grain boundaries in bcc Fe: Application of local energy and local stress. J. Mater. Sci. 2014, 49, 3980–3995. [Google Scholar] [CrossRef] [Scilit]
- Wang, J.L.; Madsen, H.K.G.; Drautz, R. Grain boundaries in bcc-Fe: A density-functional theory and tight-binding study. Model. Simul. Mater. Sci. Eng. 2018, 26, 025008. [Google Scholar] [CrossRef] [Scilit]
- Guo, X.; Zhang, J.Y.; Liu, D. Influence of alloying elements on the stability and segregation behavior of fcc-Fe/NbN interface by simulation. Vacuum 2024, 228, 113479. [Google Scholar] [CrossRef] [Scilit]
- Monkhorst, H.J.; Pack, J.D. Special points for Brillouin-zone integrations. Phys. Rev. B 1976, 13, 5188–5192. [Google Scholar] [CrossRef] [Scilit]
- Wang, H.Y.; Gao, X.Y.; Xing, L. Segregation mechanism of alloying elements at the fcc-Fe/bcc-Fe interface and its effects on carbon diffusion across the boundary. J. Phys. Chem. Solids 2023, 183, 111657. [Google Scholar] [CrossRef] [Scilit]
- Du, Y.A.; Ismer, L.; Rogal, J. First-principles study on the interaction of H interstitials with grain boundaries in α- and γ-Fe. Phys. Rev. B 2011, 84, 144121. [Google Scholar] [CrossRef] [Scilit]
- Scheiber, D.; Pippan, R.; Puschnig, L.; Romaner, L. Ab initio search for cohesion-enhancing impurity elements at grain boundaries in molybdenum and tungsten. Model. Simul. Mater. Sci. Eng. 2016, 24, 085009. [Google Scholar] [CrossRef] [Scilit]
- Rice, J.R.; Wang, J.S. Embrittlement of interfaces by solute segregation. Mater. Sci. Eng. A 1989, 107, 23–40. [Google Scholar] [CrossRef] [Scilit]
- Wu, X.B.; You, Y.M.; Kong, X.S. First-principles determination of grain boundary strengthening in tungsten: Dependence on grain boundary structure and metallic radius of solute. Acta Mater. 2016, 120, 315–326. [Google Scholar] [CrossRef] [Scilit]

Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2026 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license.