


Abstract
The performance and mechanism of heterogeneous catalytic reactions are fundamentally governed by the formation, stability, and reactivity of transient surface intermediates. These species—such as isocyanates, alkyl groups, carboxylates, formates, carbonates, alkoxy and acyl intermediates—often exist at low concentrations and with short lifetimes, making their identification challenging. This review summarizes the current knowledge on the formation, spectroscopic identification, and thermal behavior of these intermediates on metal single crystals, metal nanoparticles, and oxide-supported catalysts. Emphasis is placed on key reactions including CO and NO oxidation–reduction, CO and CO2 hydrogenation, Fischer–Tropsch-related pathways, and reforming of ethanol. Advanced surface-sensitive techniques (TDS, XPS, UPS, IR, HREELS) are highlighted for their role in elucidating intermediate structures and reaction pathways. The isocyanate surface complex is an existing intermediate in NO reduction with CO, and NCO is responsible for NH3 formation. Alkyl groups can be prepared from thermal- or photo-induced dissociation of alkyl halogenide. Oxygen-containing intermediates relevant to CO2 hydrogenation are addressed, with particular attention to formate, carboxylate, and related species. M/CeO2 (M = Pt, Rh, Ir, Ru) seems to be the best catalyst for hydrogen production from ethanol reforming. The nature of support may affect hydrogen production. The review also discusses how metal–support interactions, particle size, and surface morphology influence intermediate stability and catalytic selectivity. Overall, the work provides a comprehensive framework for understanding how transient surface complexes control technologically important catalytic transformations.
1. Introduction
Heterogeneous catalysis offers a wide range of possibilities and enables numerous technologically important reactions related to the production of fuels (including hydrogen) and fine chemicals, and the mitigation of environmental pollution. Within this context, several key catalytic reactions have been extensively investigated, including CO oxidation, NO reduction, CO hydrogenation (including Fischer–Tropsch synthesis), CO2 hydrogenation and methanation, and the reforming of methane and light alcohols. These reactions can be grouped into three main categories: (i) oxidation–reduction reactions (CO, NO); (ii) alkyl group production, identification and their reactions; and (iii) CO2 hydrogenation and reforming processes. These systems are considered representative examples in which the role of surface intermediates has been widely studied and where structure–reactivity relationships can be established. Owing to major advances in surface science, material characterization techniques, and computational chemistry, the mechanistic understanding of these reactions has improved significantly in recent decades [1,2,3,4,5,6,7,8,9,10,11].
The activity, selectivity, and mechanisms of catalytic reactions are governed by several interrelated factors. It is extensively documented that the electronic structure and morphology of active metal nanoparticles play a decisive role. Metals ranging from single atoms to nanoclusters and nanoparticles exhibit markedly different properties [5,6,7,8,9,10]. Sub-nanometer metal clusters (~1 nm) do not exhibit bulk-like electronic structures (e.g., a well-defined Fermi level), leading to distinct adsorption behavior and reactivity [9,11,12,13]. In addition, metal nanocrystals exposing high-index facets often show enhanced activity due to the high density of low-coordination sites such as steps, edges, and kinks. Size-dependent oxidation states can further influence reaction pathways and selectivity [14,15]. Moreover, strong metal–support interactions, as well as adsorbate-induced surface reconstruction, can significantly modify catalytic performance [16,17,18,19,20,21,22]. Despite these advances, it is widely recognized that catalytic reactions proceed through complex sequences of elementary steps involving transient surface intermediates, rather than via direct conversion of reactants into products.
These structural and electronic characteristics strongly influence how reactant molecules interact with catalytic surfaces. Rather than converting directly into final products, adsorbed species typically undergo a sequence of elementary steps, leading to the formation of transient surface intermediates. In most cases, these intermediates result from interactions between co-adsorbed species. The stability and reactivity of such surface intermediate species largely determine the overall activity and selectivity of catalytic processes [10,11,12,23,24,25,26,27,28,29,30,31,32,33,34,35,36,37,38]. Modern surface-sensitive techniques, including temperature-programmed desorption (TPD/TDS), infrared spectroscopy, X-ray photoelectron spectroscopy (XPS), ultraviolet photoelectron spectroscopy (UPS), and extended X-ray absorption fine structure (EXAFS), enable the detailed investigation of these species. In parallel, their energetics can be quantified through both experimental and computational approaches [39,40,41,42].
This review examines the chemical nature (stability and reactivity) of key catalytic intermediates (isocyanate, alkyl groups, carboxylate, formate, carbonyl, formyl, carbonates, alkoxy, aldehydes, acetyl, and acetate) with particular emphasis on their identification and characterization as transient surface complexes in technologically important catalytic reactions. The aim of this review is to provide a structured overview of the formation, identification, and reactivity of key surface intermediates across selected catalytic reactions. Emphasis is placed on identifying common mechanistic features, comparing model surfaces with supported catalysts, and highlighting the role of metal–support interactions in controlling intermediate stability and catalytic selectivity.
2. Formation and Preparation of Possible Intermediates Existing in Reaction Conditions
Investigating the surface chemical nature of reaction intermediates remains a significant challenge in heterogeneous catalysis, particularly when attempting to distinguish their behavior on well-defined metal surfaces (e.g., single crystals), metal nanoparticles, supports, and metal–oxide interfaces. Such comparative approaches are essential for establishing reliable reaction mechanisms. Accordingly, considerable effort has been devoted to studying the formation, stability, and reactivity of surface intermediates under realistic (in situ and operando) reaction conditions, using both experimental and theoretical methods.
In many cases, primary insights into surface species are obtained from thermal desorption spectroscopy (TDS). However, modern surface science techniques, especially vibrational spectroscopies (IR, DRIFTS, RAIRS, SFG, HREELS), enable the direct identification of adsorbed intermediates through their characteristic vibrational frequencies, which provide some of the most critical information for elucidating the nature of catalytic intermediates [43,44,45,46,47,48,49,50,51,52,53,54,55,56,57,58,59,60,61,62,63,64,65,66,67,68,69,70,71]. Furthermore, operando Fourier transform infrared (FTIR) spectroscopy can be combined with steady-state isotopic transient kinetic analysis (SSITKA) to obtain quantitative information on surface intermediates. This approach allows not only the identification of adsorbed species but also the determination of their surface lifetimes and relative abundances under reaction conditions [28,60,61]. Such combined methodologies have been successfully applied, for example, in CO2 hydrogenation, where both the nature of intermediates and the selectivity of the catalyst can be evaluated simultaneously [28].
The results of photoelectron spectroscopy (XPS, UPS), together with EXAFS, significantly contribute to the identification, formation and stability of surface complexes. Based on XPS and UPS measurements, the identity and surface reaction of alkyl intermediates (CH, CH2, CH3, C2H3 and C2H5) were determined from the decomposition of precursor molecules on metal and oxide surfaces. Processes such as formaldehyde formation from CH2 oxidation; selective oxygen addition to adsorbed CH2 and CH3 and their coupling reactions; preparation of the formate (HCOO) surface complex; and activated, negatively charged CO2 production on catalysts were easily followed by XPS and UPS [25,26,35,50,51,52,53,54,55,72,73,74,75,76,77,78,79,80,81,82,83,84,85,86,87,88,89]. Recent advances have further extended the capabilities of XPS, enabling not only high-resolution measurements [90], but also operation under elevated pressures in so-called near-ambient-pressure XPS (NAP-XPS) [91]. This latter development allows the investigation of surfaces in the presence of adsorbates under conditions much closer to realistic catalytic environments, and has provided valuable insights into reaction mechanisms during CO2 hydrogenation and ethanol transformation [62,65,92,93].
In many catalytic systems, key intermediates are formed on metal surfaces; however, their identification on metal single crystals and supported metal nanoparticles is not straightforward. One major difficulty arises from the inherently low surface concentration of these intermediates, which is often a consequence of their short lifetimes. Secondly, the oxide support can significantly influence the reactivity of intermediates formed on the metal, or the intermediates may migrate from the metal surface to the support [16,17,18,19,20,21,22]. To overcome these challenges, model precursor molecules are often employed as adsorbates, whose controlled dissociation yields the desired surface species. For example, HNCO dissociates to form isocyanate (NCO) species [23,48], formic acid (HCOOH) decomposition produces surface formate (HCOO) species [36,83,84], and the dissociation of light alcohols leads to the formation of alkoxy intermediates [37,38,70,71]. Similarly, alkyl groups can be generated by the decomposition of alkyl halides [72,73,74]. In addition, photo-induced processes can significantly facilitate the formation of surface intermediates. Under low-temperature conditions, these methods can markedly increase the surface concentration of reactive species. This approach is particularly effective for the generation of alkyl fragments via the photo-induced dissociation of photoactive alkyl halides [73].
3. Catalytic Intermediates Formed on Metal Single-Crystal Surfaces, Metal Nanoparticles and Oxide-Supported Metal Nanoparticles
Surface intermediates play a central role in determining the activity, selectivity, and mechanism of heterogeneous catalytic reactions. Their formation, stability, and reactivity depend strongly on the nature of the active phase, including metal type, particle size, surface structure, and the presence of oxide supports or metal–support interfaces. In this section, structured overview of the most important classes of surface intermediates relevant to key catalytic processes were provided, including CO and NO reactions, hydrocarbon transformations, and CO2 hydrogenation. Rather than presenting individual studies in isolation, emphasis is placed on identifying general trends, comparing different catalytic systems, and highlighting common mechanistic features.
The discussion is organized into thematic subsections according to the type of intermediates and reactions involved. First, nitrogen-containing intermediates, particularly isocyanate (NCO) species, are discussed in the context of CO–NO reactions. This is followed by an overview of hydrocarbon-derived intermediates (CHx and C2Hx species) and their transformations, including hydrogenation, dehydrogenation, and oxidation pathways. Finally, oxygen-containing intermediates relevant to CO2 hydrogenation and reforming reactions are addressed, with particular attention to formate, carboxylate, and related species.
3.1. Intermediates in CO Oxidation and NO Reduction
3.1.1. CO + O2 Reaction
Carbon monoxide oxidation is a key process in environmental catalysis. Therefore, the CO + O2 reaction has been extensively studied both theoretically and experimentally on model systems and supported catalysts [94,95]. Since the pioneering work of Langmuir [96], two fundamental pathways have been proposed: (i) a mechanism in which chemisorbed CO reacts with lattice oxygen (Oa), and (ii) a pathway in which adsorbed CO reacts with adsorbed oxygen species on the surface. The Eley–Rideal mechanism [97] may also operate, in which one reactant is adsorbed on the catalyst surface while the second reactant reacts directly from the gas phase. In this simple reaction, the most important key elementary step is the adsorption and activation of CO molecules in different adsorption mode on the catalyst surface. Geminal, linear and bridge-bonded CO can be distinguished based on their vibrational positions, which have different activities. A geminal structure (2098–2027 cm−1) appears in very-high-dispersity metals (<2 nm), and linear (2040–2060 cm−1) and bridge-bonded CO is typical in medium- and larger-sized particles (>3 nm) [10,20,21]. The oxidation state of metals also influences the IR position of CO [20]. Besides the importance of the adsorption mode of CO in its oxidation, the structure of adsorbed CO strongly depends on the oxidation state and morphology of metals on oxide surfaces; therefore, CO can be used as probe molecule in the characterization of metal nanoparticles.
3.1.2. The Role of the NCO Intermediate in NO Reduction with CO on Metals
The oxidation of CO by NO is another important reaction. The mechanism of the CO + NO reaction was also studied on metals, including single-crystal surfaces [98,99,100,101,102,103,104,105]. This reaction can proceed via two main global pathways: one leading to the desired product N2, and another resulting in the formation of the undesired by-product N2O [98]. It is accepted that both reactants (CO, NO) must first adsorb on the catalytic surface, followed by a series of elementary reaction steps [98]:
CO(g) → CO(ads)
NO(g) → NO(ads)
NO(ads) → N(ads) + O(ads)
NO(ads) + N(ads) → N2 + O(ads) (δ − N2)
2N(ads) → N2 (β − N2)
NO(ads) + N(ads) → N2O
CO(ads) + O(ads) → CO2
It should be noted that the above set of reactions is not complete. There is possibility that the adsorbed nitrogen atom can react with CO to form isocyanate (NCO) species. However, experimental evidence for this intermediate is limited, with only a few reported observations. The formation of isocyanate species (characterized by infrared vibrational frequencies at ~2240 cm−1) was tentatively identified by infrared reflection–absorption spectroscopy (RAIRS) on Pt(100) and Pd(111) [101,102]. While no clear experimental evidence for NCO formation has been reported on other metal surfaces [103,104], its preferential formation on Pd(100) and Pd(111) surfaces has been supported by theoretical studies [105]. Theoretical calculations have also shown that the reaction of CO with NO to form adsorbed NCO and CNO species may occur on Cu2O(111), with NCO formation being both thermodynamically and kinetically more favorable than CNO formation [106]. Furthermore, it has been clearly demonstrated that isocyanate surface complexes are genuine reaction intermediates on many oxide-supported metal catalysts [43,44,45,46,47,107,108,109,110]. In these systems, NCO species can be formed via the reaction of adsorbed atomic nitrogen (originating from NO dissociation) with either gas-phase or adsorbed CO molecules:
N(ads) + CO → NCO(ads)
To study the formation and thermal stability of NCO experimentally, the surface dissociation of the HNCO precursor was applied on several single-crystal metal surfaces, including Cu(111) [111], Pt(110), Pt(111) [112,113], Rh(111) [48,114], Pd(100) [23], Pd(111) [102] and Au(111) [115]. These studies consistently show that, depending on the metal, NCO species are stable only below room temperature. Upon heating, NCO decomposes into CO and adsorbed nitrogen, with decomposition typically initiating around 170 K, as evidenced by TDS and electron spectroscopies (AES, ELS). The presence of pre-adsorbed oxygen leads to a moderate stabilization of NCO species [23,108,111,113,114]. Isocyanate species could also be produced via oxidation of CN group on Cu(111) and Rh(111) surfaces [116,117,118]. Pre-adsorbed oxygen results in an increase in the relative amount of NCO, shifts the dominant vibrational band of NCO, and significantly stabilizes the NCO species on metals.
Moreover, vibration techniques (RAIRS and HREELS) offer a possibility to follow the temperature evolution of the isocyanate surface species in detail. For demonstration, the thermal behavior of NCO on Rh(111), Pd(100) and Au(111) surfaces is presented in Figure 1, Figure 2 and Figure 3. At 95 K, the strong band at 2277 cm−1 is detected due to the asymmetric stretch of molecularly adsorbed HNCO on Rh(111) (Figure 1) [48]. Upon heating to ~175 K, this species is removed (via desorption and dissociation), while a band at 2160 cm−1 appears, indicating the formation of NCO. With a further increase in temperature, the intensity of this band decreases and disappears at 300–330 K, accompanied by the formation of linearly adsorbed CO at 2070 cm−1.
Figure 1.
RAIR spectrum of adsorbed HNCO on Rh(111) and heated to higher temperature. (Reproduced from [48]).
Figure 2.
(A,B) RAIR spectra of adsorbed HNCO on Pd(100) surface and effect of heating. (Reproduced from [23]).
Figure 3.
Effects of HNCO exposure on the HREEL spectra of untreated Au(111) at ~100 K (A), and subsequent annealing (B) (exposure of HNCO was 8.0 L). (Reproduced from [115]).
A similar behavior was observed on Pd(100) (Figure 2) [23]. At low HNCO exposure (6.0 L), the dominant absorption band at 2274 cm−1 is assigned to molecularly adsorbed HNCO. The inset of Figure 2 shows the N–H stretching region, with bands at 3374 and 3240 cm−1. The coverage-dependent NCO band for a(NCO) (2215–2147 cm−1) was detected in the 160–280 K temperature range. From ~180 K, another peak evolved at 1910–1928 cm−1, attributed to bridge-bonded CO formed via NCO decomposition.
Representative HREELS spectra of Au(111) recorded at ~100 K under different levels of HNCO exposure are shown in Figure 3A. The adsorption of HNCO gives rise to several vibrational losses at 515, 900, 990, 1390, 2270, 2780, and 3230 cm−1 [115]. The most prominent feature is the loss at the 2270 cm−1, attributed to the asymmetric stretching of molecularly adsorbed HNCO (νa(NCO)). Additional losses at 2780 and 3230 cm−1 are assigned to N–H stretching modes, while the adsorption features of the NCO group align with the literature data obtained previously [119]. After heating the adsorbed layer, the attenuation of vibration loss at 2270 cm−1 was experienced, and a new spectral feature at 2185 cm−1 at around 150 K, assigned to the asymmetric stretching mode of NCO, was observed (Figure 3B). This band, together with the symmetric stretching mode of NCO (νa(NCO)) at ~1390 cm−1, remains detectable up to ~430 K, indicating a higher thermal stability of NCO on Au(111) compared to that on single-crystal surfaces of Pt-group metals, like Pd and Rh. At the same time, no vibrational features corresponding to adsorbed CO were detected, as CO does not adsorb on clean Au(111) surfaces under these conditions.
HNCO(ads) → NCO(ads) + H(ads)
NCO(ads) → CO(g) + N(ads)
Although it can be clearly seen that NCO is instable with clean metal, a marked difference is that its decomposition depends on the nature of the metal. It disappeared at 285 K on Pd(100), and this value was 430 K on Au(111). The differences in stability correlate with the affinity of the metal to CO adsorption. The adsorption bond of CO on Rh(111) and Pd(100) is high, while the CO bonds weakly to Au(111).
3.1.3. Identification of NCO on Metal Nanoparticles on Oxides
In contrast to clean, unsupported metal surfaces, the isocyanate exhibits substantially higher stability on oxide-supported metal catalysts, persisting under typical NO + CO reaction conditions [43,44,45,46,47,107,108,109,110,120,121,122]. Isocyanate intermediates were first identified by means of infrared spectroscopy by Unland on supported noble metals [43]. He proposed that the NCO intermediate could be responsible for the formation of NH3 in the presence of water at 400 °C during the NO + CO reaction.
Subsequent studies have systematically investigated the formation, stability, and reactivity of NCO species on metals on various oxide catalysts. Figure 4A illustrates the formation and thermal stability of NCO species on Pt/Al2O3 formed during NO + CO reaction [45]. Infrared bands corresponding to adsorbed NO appear in the 1500–1650 cm−1 range, and linearly adsorbed CO was found at 2080 cm−1. In strong alignment with the findings of Unland [43], a characteristic va(NCO) mode of NCO at 2267 cm−1 could be clearly identified.
Figure 4.
Infrared spectra observed at room temperature after dosing 5% Pt/Al2O3 sample with a 10% CO, 5% NO and 85% N2 mixture at different temperatures for 1 min (A). Dependence of the intensities of the band at 2267 cm−1 (observed at room temperature) (B). (Reproduced from [45]).
e temperature dependence of NCO formation is shown in Figure 4B. This band appears at ~150 °C, reaches a maximum around 400 °C, and then decreases, becoming barely detectable near ~500 °C. This behavior suggests that partially reduced metal sites (e.g., Pt) play a crucial role in activating NO molecules, promoting their dissociation, and facilitating the reaction between CO and adsorbed nitrogen atoms (Equation (8)).
Recent qualitative measurements indicate that the number of adsorbed NCO species can exceed the number of surface Pt atoms [43,46], suggesting that a fraction of NCO species is located on the alumina support [46,108]. This hypothesis is further supported by the observation that NCO can also produce on non-metallic chromia/alumina catalysts [121]. It has also been demonstrated that the nature of the support strongly affects the vibrational frequency of NCO due to the strong metal interactions (SMSIs). For Pt catalysts of comparable particle size and under identical NO + CO reaction conditions, the νₐ(NCO) band is observed at 2210 cm−1 on Pt/TiO2, 2241 cm−1 on Pt/MgO, 2272 cm−1 on Pt/Al2O3 and 2318 cm−1 on Pt/SiO2 [110]. The observed IR frequences are provided in Table 1. While the thermal stability of NCO on supported Pt varies with the support, no NCO bands are observed in the absence of Pt, indicating that metal sites are essential for its formation. The stability of NCO increases in the following order:
Pt/TiO2 < Pt/Al2O3 < Pt/MgO < Pt/SiO2
Table 1.
The positions of the bands are due to surface isocyanate and carbon monoxide (data are reproduced from [110]).
Taken together, evidence from single-crystal metals, metal nanoparticles, and supported metal catalysts suggests the following mechanism: NCO initially forms on the metal surface, but it is highly unstable there. Therefore, it subsequently migrates to the metal/support interface or onto the support itself. In this way, NCO acts as an effective intermediate in the NO + CO reaction, capable of reacting with adsorbed or gas-phase NO. The efficiency of this step depends on the stability of NCO at the interface.
NCO(ads) + NO → CO2 + N2
We note here that NCO can form stable compounds, especially when the NCO is located on the support. In these cases, the NCO can react with the cation of oxide forming compounds as a side product, for example Si(NCO)4. Moreover, NCO species have been implicated in the formation of trace CN-containing products [111,112,113,114,115,116,117,118,119,120,121,122], and it has been proposed that NCO intermediates can contribute to NH3 formation in the presence of water under conditions relevant to automobile exhaust [43,44,45,46,107]. Recent studies have experimentally confirmed that NH3 can indeed form from NCO intermediates [109].
NCO(ads) → NC(ads) + O(ads)
2 NC(ads) → C2N2(g)
NCO(ads) + H(ads) + H2O → NH3 + CO2
3.2. Identification and Reactivity of Alkyl Groups, Fischer–Tropsch Synthesis, CO Hydrogenation
Alkyl groups (CH, CH2, CH3, C2H3, C2H5, etc.) are key intermediates in Fischer–Tropsch synthesis and in the catalytic conversion of hydrocarbons into oxygenated products such as alcohols [47,52,72,73,81,123,124,125,126,127,128,129]; therefore, understanding their chemistry is crucial. In Fischer–Tropsch chemistry, two principal mechanistic pathways are generally considered. In the first pathway, the C–O bond of adsorbed CO (or CO-derived species such as C–OH) is cleaved, leading to the formation of CHx species that participate in chain growth. In the second pathway, CO insertion into a growing hydrocarbon chain occurs, resulting in the formation of oxygenated intermediates of the type R–CHx–OH [129]. A detailed understanding of these mechanisms requires the identification and characterization of the transient surface intermediates involved, particularly CHx and CxHy species.
3.2.1. Identification and Dehydrogenation–Hydrogenation of Alkyl Groups
To address this, XPS, UPS, and various vibrational techniques (DRIFTS, RAIRS, and HREELS) have been employed to identify transient surface species and to provide valuable information on their stability and reactivity. In most cases, alkyl fragments were generated via thermal- or photo-induced decomposition of alkyl halides. Previous studies reported the identification and stability of CH3, CH2, and CH species on Co and Ni polycrystalline surfaces using XPS and UPS following the thermal dissociation of CH3Cl and CH2Cl2 [72]. The early cleavage of the C–Cl bond enabled the investigation of the dehydrogenation of CHx surface species without interference from other functional groups. Table 2 shows the photoemission (PE) energies for C 1s in XPS and the corresponding CHx features in UPS obtained on Co and Ni surfaces. The binding energies are referenced to the Fermi level of the metal. The temperature ranges over which each species is observed, and the dominant species are also indicated. The facile cleavage of the C–Cl bond enables the study of the stepwise dehydrogenation of CHx surface species in the absence of significant interface from the support (Equation (15). Complementary TDS measurements indicate that a fraction of the adsorbed hydrogen recombines to form H2, while another fraction reacts with CH3 and CH2 species to produce methane in the gas phase (as shown in Equations (19) and (20)).
CH3Cl(ads) → CH3(ads) + Cl(ads)
CH3(ads) → CH2(ads) + H(ads)
CH2(ads) → CH(ads) + H(ads)
CH(ads) → C(ads) + H(ads)
CH2(ads) + 2H(ads) → CH4(gas)
CH3(ads) + H(ads) → CH4(gas)
Table 2.
Characteristic PE line energies for adsorbed on polycrystalline Co and Ni.
The vibrational frequencies and assignments of alkyl intermediates have been reported for a wide range of metal surfaces, including Ag, Cu, Ni, Fe, Pt, Rh, and Ru, and are summarized in Table 3. In most studies, alkyl species were generated via thermal- or photon-induced decomposition of alkyl halides, typically adsorbed at low temperatures (90–100 K). Notably, the vibrational frequencies of alkyl species exhibit only a weak dependence on the nature of the metal surface, indicating that their bonding characteristics are largely preserved across different metals.
Table 3.
Vibrational characteristics of alkyl species on clean metal (Ag, Cu, Fe, Pt, Rh, Ru) surfaces.
The CH3 species on most metals can be characterized by the asymmetric and symmetric stretching modes (νa(CH3)), νs(CH3)) typically observed at 2918–2950 cm−1 and 2775–2821 cm−1, as well as the asymmetric and symmetric deformation modes (δa(CH3)), (δs(CH3)), which usually appear at 1350–1435 cm−1 and at 1141–1185 cm−1 (Table 3). Figure 5 shows the RAIR spectrum of CH3 on a clean Rh(111) and on potassium-, iodine- and zinc-modified Rh(111) surfaces. In these experiments, the methyl radicals were generated by direct decomposition of azomethane (CH3N2CH3) in front of the sample [49]. Three significant vibrations (νa(CH3), δa(CH3), (δs(CH3)) were observed on the clean sample (Figure 5A). The symmetric band of CH3 cannot be detected by RAIRS due to dipole selection rules.
Figure 5.
(A) RAIR spectra of the adsorbed CH3 at saturation on clean and doped Rh(111) at 100 K. (B) Absorbance of the peak of CH3 at 2918–2926 cm−1 after heating the adsorbed layer on clean and doped Rh(111). The coverage of I and Zn was ca. 0.3. Reproduced from [49].
The thermal stability of CH3 is illustrated in Figure 5B, which shows the temperature-dependent evolution of methyl species at 2918–2926 cm−1 on clean and modified Rh(111) surfaces. This band disappears completely at 300–320 K, indicating the removal or transformation of CH3 species.
Consistent results have been obtained by HREELS, where CH3 species were generated via thermal- or photo-induced dissociation of alkyl iodides or methane. Overall, the thermal stability of CH3 on Rh surfaces is comparable to that observed on other clean metal surfaces [53,73,74,125,130,131,132,133,135,136,139]. Figure 5B further demonstrates that potassium significantly enhances the stability of CH3, whereas Zn and I induce only minor stabilization effects. In contrast, co-adsorbed CO does not affect either the vibrational frequencies or the thermal stability of CH3 species [135]. The influence of oxygen is more complex and will be discussed separately.
In Fischer–Tropsch synthesis, hydrogenation–dehydrogenation reactions of alkyl species play a critical role. As observed by Co and Ni, TDS studies confirm that both these pathways occur on noble metals, leading to the formation of CH2, CH, C, and CH4 [125]. Notably, the coupling reactions of CH3 species were not observed under these conditions.
The identification, stability, and reactivity of CH2 species have also been extensively investigated on metal surfaces using a combination of spectroscopic techniques (XPS, UPS, RAIRS, HREELS) and mass spectrometry [50,51,52,53,135,136,137]. Figure 6 illustrates the general features of CH2 surface chemistry on Pt(111), providing a representative example applicable to other metal surfaces. In this case, chloroiodomethane (ClCH2I) was used as a precursor due to its favorable dissociation characteristics [50,51]. At 100 K, the monolayer of ClCH2I exhibits losses at 500, 725, 1140, 1365, 2955, and 3020 cm−1. In harmony with TPD and XPS results, the C–halogen bond cleavage occurs in the temperature range of 170–200 K. Subsequently, CH2 species identified by their symmetric, asymmetric and deformation modes at 1440, 2880 and 2940 cm−1 appear above ~230 K [50,51]. These bands disappear around 320 K; the higher-frequency band already contains contributions from CH species. Isotope exchange experiments indicated the formation of methane in hydrogenation reactions of surface CH2. In addition, minor halogen-assisted CCH formation has been detected around ~540 K, attributed to the reaction between adsorbed C and CH species, with a characteristic band at 3035 cm−1 [54].
Figure 6.
HREEL spectra for monolayers and multilayers of ClCH2I at 100 K on Pt(111) and for multilayers warmed up briefly to various temperatures as indicated. All spectra were taken at 100 K. Reproduced from [50].
In contrast to Pt and Rh surfaces, the CH2 species on Ru(001) and Pd(100) not only undergo hydrogenation to form CH4, but also readily dimerize to produce C2H4 in the temperature range 160–230 K, as demonstrated by XPS, UPS, and TDS measurements [81,82]. A fraction of the formed ethylene, adsorbed in a π-bonded configuration, desorbs, while the remaining fraction, adsorbed in a di-σ configuration, is further transformed into vinyl and ethylidyne species between 220 and 300 K; these species only decompose above 450 K. On Cu surfaces, CH2 species similarly react to form C2H4 at 160–230 K, which desorbs with a peak temperature (Tp) of 240 K. In contrast to Pt-group metals, no self-hydrogenation of CH2 to CH4 is observed on Cu(100) [71]. On Ag(111), CH2(a) couples with neighboring CH2(a) to form ethylene, which desorbs in a reaction-limited process at 230–260 K, as evidenced by TPD and RAIRS experiments [130].
The formation and reactivity of adsorbed C2H5 species on Rh(111) was studied by means of RAIRS and TDS. In these experiments, hydrocarbon fragments were also produced via thermal- and photo-induced dissociation of corresponding iodine compounds. The conclusions drawn for Rh(111) are also generally applicable to other Pt metals (see Table 3). Figure 7A shows spectra for the clean surface, while Figure 7B illustrates the effect of potassium modification. At 100 K, the observed vibrational features correspond to the gas-phase and adsorbed C2H5I, consistent with data reported for Pt(111) [138]. During annealing, C–I bond cleavage occurs at ~140 K on clean Rh(111), and by 200 K, the spectra exhibit bands that are characteristic exclusively of adsorbed C2H5 species (see the assignments in Table 3). Further heating to ~240 K leads to the disappearance of C2H5 features and the emergence of new bands at 2975, 2879, 1380, and 1125 cm−1, which are characteristic of ethylidyne species, likely formed via intermediate vinyl species [66]. Simultaneously, dehydrogenation produces C2H4, which desorbs at Tp = 185 K, while a fraction of the species undergoes hydrogenation to form C2H6 at similar temperatures. At higher temperatures (>330 K), a band at ~3030 cm−1 indicates the formation of stable CCH species.
Figure 7.
Effects of temperature on the RAIR spectra of adsorbed C2H5I on Rh(111). (A) Clean Rh(111); (B) K-dosed Rh(111), ӨK = 0.36. In the latter case, potassium was deposited on the adsorbed C2H5I heated to 190 K. (Reproduced from [53].
The presence of potassium significantly alters the reaction pathway, and promotes the direct formation of adsorbed C2H4 (bands at 2947, 2912, 1195, and 1117 cm−1) at ~190 K, which subsequently desorbs at higher temperatures. In addition, potassium induces coupling reactions of ethyl species, leading to the formation of butene and butane (see Equation (26)). Similar reaction pathways have been reported for other clean Pt-group metals [53,55,73,75,76,138]. In general, C2H5 species transform to yield C2H4 and C2H6 at 160–240 K. A fraction of the resulting C2H4 is further converted into vinyl species, which remain stable up to ~450 K (Equations (23)–(25)).
C2H5(ads) → C2H4(ads) + H(ads)
C2H4(ads) → C2H4(g)
C2H4(ads) → CHCH3(ads)
CHCH3(ads) → CCH3(ads) + H(ads)
CCH3(ads) → CCH(ads) + 2H(ads)
2C2H5(ads) → C4H10(g)
The same steps were observed on Ni surfaces; however, the dehydrogenation proceeded much faster [140]. Cu surfaces exhibited comparable behavior in C2H5 transformation, although with a stronger dependence on surface orientation (facet selectivity) [80].
3.2.2. Interaction of Alkyl Groups with Adsorbed Oxygen on Metals
CH3 + oxygen reaction
The interaction of alkyl groups with adsorbed oxygen, leading to the formation of alkoxide species (e.g., CH3O, CH2O, C2H5O), provides important insight into the formation of oxygenated products, such as aldehydes and alcohols, during Fischer–Tropsch synthesis [140,141]. Among these, the interaction between methyl species and adsorbed oxygen has been studied detail, as it represents the simplest model system for understanding alkoxide formation. This interaction has been investigated primarily by RAIRS, in which methyl groups were generated in situ via the decomposition of azomethane, thereby avoiding the influence of halogen-containing precursors [49].
As discussed previously, the characteristic vibrational bands of adsorbed CH3 are observed at 2918 cm−1 (νa(CH3)), 1353 cm−1 (δa(CH3)) and 1141 cm−1 (δs(CH3). These features are also present when CH3 is adsorbed on oxygen-covered Rh(111) surfaces (Figure 8A). addition, new bands appear at 2960, 1440 and 1040 cm−1, which are assigned to methoxide (CH3O) species, corresponding to νa(CHa), δs(CH3) and ν(C-O) vibrational modes, respectively [52,140,141,142,143,144]. In The formation of CH3O species has also been confirmed by HREELS measurements [47]. Methoxide is a key intermediate in the synthesis of alcohols and in reactions such as CH4 + CO2 conversion [28,33,62,64]. As shown in Figure 8A,B, CH3O species can form both when CH3 is adsorbed on an oxygen-covered Rh(111) surface and when oxygen is introduced onto a CH3-saturated surface. Upon heating the co-adsorbed layer, decomposition begins at ~158 K, leading to the formation of adsorbed CO. The methoxide intermediate remains stable up to 188–220 K (Figure 8). Subsequently, adsorbed CO and hydrogen (produced via dehydrogenation of CHx species) react with oxygen to form CO2 and H2O.
Figure 8.
RAIR spectra of Rh(111) (A) following the adsorption of CH3 on O-covered surface (ϴO = 0.5), and (B) following the adsorption of oxygen on the CH3-saturated surface. Both compounds were adsorbed at 100 K. (Reproduced from [49]).
CH2 + oxygen reaction
A general feature observed across metal surfaces is that pre-adsorbed oxygen slightly suppresses the dissociation of the C–halogen bond in CH2I2, while simultaneously strengthening the interaction of the resulting CH2 species with the surface. In addition, increasing oxygen coverage leads to a decrease in the formation of CH4 and H2, which are typical products in the absence of oxygen. A key outcome of the interaction between CH2 and adsorbed oxygen is the formation of formaldehyde (CH2O), the surface concentration of which depends strongly on the nature of the metal. On Pt-group metals and several other surfaces, CH2 reacts with adsorbed oxygen above ~170 K to form CH2O; however, this species desorbs rapidly after formation in a rate-limiting step, making its direct spectroscopic detection difficult [50,51,55,78,79,80,81,82,85,130]. Consequently, TPD provides more detailed insight into its formation and subsequent transformation:
CH2(ads) + O(ads) → CH2O(ads)
CH2O(ads) → CH2O(g)
CH2O(ads) → -CO(ads) + 2H(ads)
CH2O(ads) + O(ads) → -OCHO-(ads) + H(ads)
-OCHO-(ads) → CO2(g) + H(ads)
CO(ads) + O(ads) → CO2(g)
2H(ads) + O(ads) → H2O(g)
Ru is an excellent catalyst for the synthesis of higher hydrocarbons; therefore, it was interesting to examine its effect on the coupling of CH2 with O [81]. XPS measurements indicate that C–I bond cleavage in CH2I2 occurs around 200 K, leading to the formation of CH2 species that react readily with adsorbed oxygen; a significant fraction of the resulting CH2O desorbs immediately in a rate-limiting step (Equations (28) [81]. Similar behavior has been reported for Pt(111), Rh(111), and Pd(100) [50,51,82].
However, the subsequent transformation of formaldehyde on Ru(001) is more complex than on other metals. TDS measurements at an oxygen coverage of ϴO = 0.25 reveal a rich reaction network (Figure 9). Compared to the clean surface, molecular desorption of CH2I2 occurs at higher temperatures, while CH4 formation at 220–230 K indicates that partial self-hydrogenation of CH2 still takes place. Smaller amounts of H2 are also detected (Tp ≈ 270 and 490 K). The most significant feature is the formation of CH2O (Tp = 298 K), confirming its role as a primary intermediate. At higher temperatures (>500 K), CO, CO2, and H2O appear as final decomposition and oxidation products.
Figure 9.
Thermal desorption spectra for various products at O = 0.25. CH2I2 exposure was 6 L. Dotted curves represent TPD products that form a clean surface. (Reproduced from [81]).
UPS further reveals the nature and evolution of surface intermediates (Figure 10). At 110 K, the spectra showed the molecular orbitals of adsorbed CH2I2 on Ru(001), which were also previously detected on Pd(100) and Cu(100) [78,79,82]. Upon heating to 219 K, new photoemissions were experienced at 5.7, 8, 8, and 11.0 eV due to adsorbed CH2O [77]. This spectroscopic observation indicates that CH2O can remain adsorbed on the Ru(001) surface, like its behavior on Cu(110).
Figure 10.
UPS spectra of Ru(001) as a function of annealing temperature at ϴO = 0.5. CH2I2 exposure was 6 L. (Reproduced from [81]).
In contrast, formaldehyde species did not remain on Pt(111), Rh(111) and Pd(100) [50,51,82]. At 298 K, additional spectral features (5.0 8.5, 9.0 and 10.7 eV) appear, which can be attributed to adsorbed formate (HCOO) species [36,77,83,84]. Formate formation has also been suggested on Ag(111) based on RAIRS measurements [134]. At temperatures above ~450 K, peaks characteristic of adsorbed CO (8.4 and 10.9 eV) and water (6.2, 8.5, and 12.3 eV) are observed [84,85,86]. Based on these findings, the following additional steps are proposed to occur on oxygen-covered Ru(001):
CH2O(ads) + O(a) → HCOO(ads) + H(ads)
2HCOO(adsz) → H2O(ads) + CO(ads) + CO2(g)
CO(a) → CO(g)
CxHy(a) + O(ads) → CO(g) + CO2(g) + H2(g)
Independent studies of formaldehyde adsorption on Ru(001) confirm that its thermal stability, decomposition pathways, and formate (HCOO) formation are consistent with those observed during CH2 + O reactions. Moreover, formate species have been directly identified by HREELS, and infrared studies demonstrate that adsorbed formaldehyde can be converted to formate on oxide-supported metal catalysts [144,145,146,147,148,149].
C2H5 + oxygen reaction
C2H5 surface groups and their interaction with oxygen also play an important role in the formation of oxygenated hydrocarbons. The oxidation of C2H5 has been studied on Rh(111) and Ru(001) surfaces [52,53,55,76,148]. In contrast to CH3 species, transiently formed ethoxy (C2H5O) is hardly detectable experimentally by spectroscopic methods. This is most likely the case due to its limited thermal stability; for example, ethoxy species generated via ethanol decomposition on clean Rh(111) decompose completely below 250 K, yielding adsorbed hydrogen and CO [148]. Upon heating, the interaction of adsorbed C2H5 with oxygen leads to the formation of H2O, CO2, and small amounts of CH3CHO and CO. The formation of acetaldehyde is generally attributed to β-hydrogen elimination of adsorbed ethoxy species, as established by Houtman and Barteau [69]. Spectroscopic measurements only reveal a loss in the CO stretching region (2040–2050 cm−1) when the T ≥ 2070 K, suggesting that oxygen-containing C2 intermediates are present only in low concentrations on Rh(111).
On Ru(001), significantly different oxidation products were experienced during C2H5 oxidation [76]. The presence of adsorbed oxygen suppressed the formation of ethane and ethylene, which were dominant on clean Ru(001), and instead promotes the formation of oxygenated products. A notable feature of this system is the strong dependence of product distribution on oxygen coverage, as evidenced by TPD-MS measurements (Figure 11). At low oxygen coverage, diethyl ether is formed, likely via coupling of two ethyl species; the involvement of short-lived ethoxy intermediates in this process cannot be excluded. At higher oxygen coverages, acetaldehyde becomes the dominant product, consistent with hydrogen abstraction from transient ethoxy species.
Figure 11.
TPD spectra of (A) acetaldehyde and diethyl ether (B) as a function of oxygen coverage. The exposure of C2H5I was 3.6 L. (Reproduced from [55]).
This pathway is analogous to the ethanol decomposition mechanisms reported for Pt-, Rh-, and Ir-based systems [37,38]. CO formation is also observed, and may originate from the oxidation of CCH surface species, which are likely formed via ethylene transformation on oxygen-free sites (steps 9–11).
C2H5(ads) + O(ads) → C2H5O(ads)
2C2H5O(ads) → C2H5OC2H5(g)
C2H5O(ads) → CH3CHO(g) + H (ads)
Thus, the overall reaction network on Ru(001) reflects a competition between hydrocarbon coupling and oxidation pathways, strongly modulated by oxygen coverage (Scheme 1).
Scheme 1.
Representation of formation of diethyl ether and acetaldehyde on oxygen-covered Ru(001).
It is worth mentioning that the formation of C–O bonds is not limited to ethyl species; similar behavior has been reported for higher alkyl groups. For instance, propoxide formation has been identified by XPS and RAIRS following thermal activation of 2-propyl groups in the presence of chemisorbed oxygen on Ni(100) [150].
In contrast to metal surfaces, alkyl species are more readily stabilized on oxide supports, where they preferentially bind to surface oxygen atoms to form alkoxy or formyl, formaldehyde (H2CO), ether or acetaldehyde species. The stability and reactivity of these intermediates are primarily governed by the nature of oxides and the characteristics of the metal–oxide interface. Both the bonding strength and coordination mode of alkoxy species play a decisive role in determining the pathways leading to oxygenated hydrocarbons on supported catalysts. Alkyl species bond to oxides, or more frequently to vacancies of oxides, and the atomic hydrogen is mostly supplied by the metal for further reaction. The formation, and reactivity of alkyl intermediates on metal nanoparticles supported by oxides are highly structure-sensitive processes [4,6,9,11,16,150,151]. This structure-sensitive process is perturbed by SMSI in Fischer–Tropsch reactions [152]. As the metal–support interaction influences the particle size, oxidation state, bulk-surface composition and dispersion, the stability and reactivity of alkyl groups depend on the strength of SMSI in different metal–oxide systems, which changes the metal structure. The Fischer–Tropsch synthesis is structure-sensitive but support-insensitive. Industrially, Co nanoparticles are commonly supported on oxides, the precise calculation was made for Co supported by different oxides. The increasing cobalt−support interaction strength trend of MgO > SiO2 > Al2O3 > TiO2 > CeO2 is in line with the results in the literature.
3.3. CO2 Hydrogenation and Methanation on Metals and Supported Catalysts
One of the most promising reactions of CO2 conversion is the CO formation via reverse water–gas shift (RWGS) reaction. The other important reaction in CO2 hydrogenation is methane formation (called the Sabatier reaction) [24,57]. Another effective method for CO2 utilization is methanol formation [24,41]. These processes often occur simultaneously, with RWGS competing with methanation and methanol formation, producing CO and H2O as by-products.
CO2 + H2 ↔ CO + H2O
CO2 + 4H2 → CH4 + 2H2O
CO2 + 3H2 → CH3OH + H2O
The catalytic conversion of CO2 is of major importance for mitigating CO2 emissions and producing value-added fuels and chemicals. Among the most relevant processes are CO2 hydrogenation and reforming reactions with hydrocarbons (e.g., CH4) [6,7,32,153,154,155,156,157,158]. Methane dry reforming reaction (CH4 + CO2 or CH4 + CO) can also serve a source of CO2 elimination with hydrogen and synthesis gas formation for energy applications [32]. The products of CO2 conversion are illustrated in Figure 12. This review primarily focuses on the identification and reactivity of intermediates formed during CO2 hydrogenation leading to CO, CH4, and light alcohols [5,6,7,127,142,143,153,154,157,158,159,160,161,162,163], as well as on hydrogen production via reforming reactions [8,32,37,38,70,71].
Figure 12.
Products from CO2 hydrogenation. (Reproduced from [154]).
3.3.1. CO2 Activation and Formate Intermediate Production on Metal Surfaces
The activation of the relatively inert CO2 molecule represents the first and most critical step in CO2 conversion reactions on both metal and oxide surfaces [25,26,27,86,87,88,89]. The formation, characterization and dissociation of electronically excited CO2− have been widely investigated on metal surfaces [86,87,88,89]. Electron donation to CO2 is typically facilitated by alkali-promoted metals or by co-adsorbed hydrogen species. At low potassium surface coverage, activated CO2 species can dissociate into adsorbed CO, which may subsequently desorb; XPS and UPS measurements indicate that this process can occur at temperatures as low as ~130 K. At higher potassium coverages, CO2− species may undergo disproportionation, yielding CO and carbonate species. In CO2 hydrogenation, carbonate species are generally considered inactive species; however, in certain structures and configurations, close to the interface, the carbonates could take part in the hydrogenation process [27,29,32,58].
CO2−(ads) → CO(ads) + O−(ads)
2CO2−(ads) → CO(ads) + CO3−(ads)
Mechanistically, CO2 hydrogenation can proceed via multiple pathways, including direct CO2 dissociation (the RWGS route) and the formate-mediated pathway [24,57]. The identification and characterization of surface intermediates are therefore essential for elucidating reaction mechanisms. Table 4 summarizes the key intermediates detected primarily by DRIFTS, including species relevant to both CO2 hydrogenation and reforming reactions. It should be emphasized that the assignment of vibrational bands to specific surface intermediates is not always straightforward, especially in the carbonate region, where overlapping spectral features, coverage effects, and metal–support interactions can lead to ambiguity and alternative interpretations. In some cases, Table 4 provides broader ranges for reported vibrational frequencies and assignments to reflect this diversity.
Table 4.
The assignments of IR bands of intermediates formed in CO2 hydrogenation, and reforming of methane and light alcohol (methanol, ethanol).
On clean metal surfaces, the intermediates involved in CO2 hydrogenation are often difficult to detect experimentally due to their low surface concentration and short lifetimes. Fisher and Bell [47] proposed that CO2 hydrogenation on Rh proceeds via mechanisms analogous to CO hydrogenation, with C–O bond cleavage as the rate-limiting step. This cleavage may occur either via direct CO dissociation or through hydrogen-assisted pathways involving HCO intermediates. Methane formation is explained by the surface hydrogenation of HxCO species.
On Pt(111), experimental investigation of CO2 hydrogenation is particularly challenging, and studies on Pt nanoparticles—especially in the absence of oxide supports –remain limited. The observed low catalytic activity is often attributed to the scarcity of low-coordination sites and weak interaction with inert supports such as SiO2 [57]. Supporting uniform Pt nanoparticles (~5 nm) on defect-free hexagonal BN enabled more controlled studies of elementary steps and the distinct activity of Pt nanoparticles could be experimentally determined [171,172]. The RWGS mechanism was established in the absence of oxide support in harmony with DFT calculations [24,57]. In the Pt/h-BN system, CO production from CO2 hydrogenation occurs via a HOCO intermediate, as evidenced by DRIFTS:
CO2(g) + H2(g) → *CO2(ads) + 2H(ads)
*CO2(ads) + H(ads) → H-O-C-O(ads)
H-O-C-O(ads) → COads + OHads
In addition to the idealized surfaces often considered in model studies, real catalysts contain a high density of surface defects (e.g., steps, edges, kinks, and vacancies), as well as residual impurities. These features can significantly influence the formation, stability, and reactivity of surface intermediates, or even introduce new reaction pathways. For instance, oxygen vacancies formed during the activation of reducible supports are highly reactive and can preferentially stabilize key reaction intermediates such as carboxylate and formate species, thereby altering reaction mechanisms and product selectivity [173]. Conversely, impurities or strongly adsorbed species may block active sites or modify the electronic structure of the catalyst, leading to either the inhibition or promotion of specific reaction steps [25].
The formate (HCOO) pathway plays a central role in CO2 hydrogenation on supported catalysts; however, formate species could not be detected on clean metal surfaces. To address this limitation, formic acid (HCOOH) is commonly used as a precursor, as it readily dissociates on metal surfaces to produce adsorbed formate species [36,83,84]. This approach also facilitates the identification of formate species on supported catalysts [30,66,154]. The HCOO species is generally unstable on metal surfaces, decomposing below ~300 K to yield adsorbed CO and gaseous CO2. Its stability can be moderately enhanced by electropositive promoters such as potassium or by co-adsorbed oxygen, while impurities such as boron can facilitate O-H bond cleavage [174]. On oxide surfaces, formic acid can also lead to the formation of formaldehyde at oxygen vacancy sites, which may subsequently decompose to adsorbed CO:
HCOOH(g) → HCOO(ads) + H(ads)
HCOO(ads) → CO2(g) + H(ads)
HCOO(ads) → CO(ads) + H(ads) + O(ads)
HCOOH(ads) → CH2O(ads) + O(ads)
CH2O(ads) → CO(g) + H(2g)
3.3.2. Intermediates in CO2 Hydrogenation on Metal-Supported Oxide Catalysts
On oxide surfaces, CO2 activation is further assisted by surface hydroxyl groups or adsorbed hydrogen, leading to the formation of bicarbonate (HCO3−), mono- and bidentate carbonate (CO32−), and carboxylate (HCO2−) species [25,56]. These intermediates can be readily identified by infrared spectroscopic techniques, as summarized in Table 4. The formation, stability, and reactivity of these intermediates are strongly influenced not only by the active metal, but also by the nature of the support, the strength of metal–support interactions, and the structure of the metal–oxide interface. Although many intermediates are formed on metal sites, their stabilization and accumulation often occur at the metal–support interface or on the support itself.
In CO2 catalytic reduction, the final product CO can form from the direct dissociation of CO2 (Equations (44) and (45)) [27,29,32,58]. The formation of methane (or other alkanes) can be rationalized by carbon–hydrogen reactions in which the carbon originates from C–O bond cleavage in CO2 or CO molecules. Methane formation may also proceed via RWGS-related pathways involving the formation and subsequent reactions of alkyl species, as discussed in the previous section. Determining the dominant mechanism for parallel reactions is very difficult in many cases. These processes often occur simultaneously; therefore, more than one intermediate play a significant role.
In the RWGS mechanism, one of the key intermediates is the carboxylate species (H-O-C-O, HCO2−) [24,57], which can be identified by DRIFTS at 1653–1631 cm−1 for va(O-C-O) and 1298–1245 cm−1 for vs(O-C-O) modes. This intermediate appears to be strongly dependent on surface morphology; it was detected on mesoporous Co3O4 but not on cubic Co3O4 [66]. This observation suggests a shift in the methanation mechanism from the RWGS pathway to direct C–O bond cleavage. Another important intermediate is hydrogen-perturbed CO (HnCO), which has been detected by DRIFTS (see Table 4), for example on NiO, CuCo, and Co3O4 [62,66,92].
This species can be hydrogenated to formyl-type intermediates (HCO), which may undergo further hydrogenation, ultimately leading to CH4:
HOCO(a) → CO(a) + OH
CO(a) + H(a) → HCO(a)
HCO(a) + H(a) → H2CO(a) + 2H(a) → H3C(a) + OH(a)
H3C(a) + H(a) → CH4(g)
Within this reaction sequence, formyl species play a key role, as supported by both thermodynamic considerations and experimental observations. On the one hand, formyl species can decompose to adsorbed CO at surface vacancies (Equation (53)); on the other hand, they can be further hydrogenated to CH4 or converted into methoxy (CH3O), which can subsequently yield CH4 in the presence of hydrogen [24,57]. This pathway may also involve methanol formation [156,157]:
H2CO(a) + 2H(a) → H2COH(a) + OH(ads)
H2COH(a) → CH2(ads) + OH(ads)
CH2(ads) + H(ads) → CH3(ads)
CH3(ads) + H(ads) → CH4(g)
H2CO(ads) + H(ads) → H3CO(ads)
H3CO(ads) + 2H(ads) → CH4(g) + OH(ads)
CH3O(ads) + H(ads) → CH3OH(g)
The literature data suggest that the RWGS pathway, via carboxylate intermediates, occurs predominantly on non-reducible oxide-supported catalysts.
A significant fraction of CO2 hydrogenation, particularly methanation, proceeds via the formate (HCOO) pathway [24]. This route is especially relevant when formate or its precursor, bicarbonate, is formed on reducible oxide supports (e.g., ceria), where hydrogen spillover is possible [28,29,30,61,62,159,162,175]. However, this does not exclude the simultaneous participation of carboxylate species in the reaction mechanism. DRIFTS-MS studies on Ru/Al2O3 have shown that formate reacts with hydrogen more rapidly than carboxylate species, while the latter may remain on the surface and transiently block CO2 adsorption sites [64]. It is interesting to emphasize here that one of the “active” intermediates could inhibit the reaction. Formate is therefore widely recognized as a key intermediate in CO2 reduction with both hydrogen and methane, and has been extensively studied (see Table 4).
It is generally accepted that direct CO2 dissociation is negligible on most metal surfaces. Instead, spectroscopic studies, including transient DRIFTS-MS, indicate that CO2 first reacts with surface hydroxyl groups on oxide supports to form bicarbonate species (va(O-C-O), vs(O-C-O) and δ(C-O-H)). Bicarbonates subsequently react with adsorbed hydrogen on metal sites to form surface formate (HCOO), which exhibits characteristic νa(O–C–O), νs(O–C–O), and δ(C–H) vibrations. The role of formate intermediates is closely related to product selectivity in CO2 hydrogenation. In catalysts favoring the reverse water–gas shift reaction, formate is often considered a transient intermediate leading to CO formation (Equations (65) and 66)), a part of which may react with hydrogen to produce CH4.
HCOO(ads) + H(ads) → HCO(ads) + OH(ads)
HCO(ads) → CO(ads) + H(ads)
CO(ads) + H(ads) → CH(ads) + O(ads)
CH(ads) + H(ads) → CH2(ads) → CH3(ads) → CH4(g)
An alternative route for formate scheme is described, containing CH4 and CH3OH formation:
HCOO(ads) + 2H(ads) → H2CO(ads) + OH(ads)
H2CO(ads) + H(ads) → H2COH(ads)
H2COH(ads) + 2H(ads → CH3OH(g) + OH(ads)
H2COH(ads) → H2C(ads) + OH (ads)
CH2(ads) + H(ads) → CH3(ads) → CH4(g)
At the same time, several studies have suggested that formate can behave as a relatively stable spectator species on certain surfaces and under specific conditions, not directly participating in the reaction and potentially poisoning active sites [176]. Therefore, controlling the formation and reactivity of formate intermediates is crucial for steering product selectivity in CO2 hydrogenation. In certain structures and configurations, close to the interface, the carbonates could take part in hydrogenation process [27,29,32,58].
The sub-nanosized range (including single-atom catalysis) is a significant part of the studies, because there is no metallic character (no Fermi level). In these cases, “molecular-like interactions” (between metal atoms or clusters and reactant molecules) dominate in the mechanisms [9,11,12,16,17]. In addition, the size-dependent electronic structure is more significant when the metal nanoclusters consist of less than 40 atoms. Metal nanoclusters formed on oxide supports have been found to be more catalytically active depending on the number of atoms forming the clusters. Strong size-dependent activity was observed in the Au/TiO2 system in CO2 hydrogenation [13]. The “molecular-like” clusters can be stabilized on several oxides. Besides small clusters, single atoms or ions (Mn+) can be stabilized by oxides, which could contribute to further progress in heterogeneous catalysis.
From a technological point of view, the activity, and mainly selectivity, are fundamental challenges in CO2 hydrogenation catalyst design. Selectivity is governed by multiple factors; however, the catalyst structure and the bonding mode and reactivity of surface intermediates appear to be particularly decisive. For example, two Co3O4 structures exhibit markedly different behavior in CO2 methanation. On the cubic structure, only carboxylate species were detected, and both CO and CH4 were formed [66]. In contrast, on mesoporous Co3O4, formate intermediates dominate, leading to nearly 100% CH4 selectivity with negligible CO formation. When both cobalt oxides are modified with small amounts of Pt, the basicity increases and the selectivity remains largely unchanged, while the activity is significantly enhanced [67].
Szanyi and co-workers demonstrated for Ni-based catalysts that, over a temperature range of 300–1100 K, selectivity shifts from high CH4 formation to predominantly CO production. This behavior is associated with the formation of carbide-like surface species, which weaken CO adsorption and favor its desorption rather than further hydrogenation. Upon regeneration of the catalyst (removal of the surface carbide), high CH4 selectivity was restored [68]. A similarly pronounced effect on selectivity has been reported for In–Pd intermetallic compounds (InPd2, InPd, and In3Pd2) [177]. Electronic interactions can change widely in In–Pd systems as a result of the bimetallic structure formation enabling tunable product distributions. Compared to monometallic Pd, InPd enhances methanol selectivity by ~70%, whereas In3Pd2 exhibits nearly 100% selectivity toward CO in CO2 hydrogenation.
3.4. Importance of Intermediates in Hydrogen Evolution Reactions
Although the highlighted subject of this part is the hydrogen production from ethanol, from the reaction-intermediate-related point of view, it is worth briefly mentioning the other two hydrogen production reactions, namely the reforming of methane and methanol.
3.4.1. CH4 + CO2 Reaction
The investigation of hydrogen sources as potential energy carriers is at the forefront of the current research. The transiently formed intermediates discussed above also play a significant role in the reforming reaction. The reaction of carbon dioxide with methane (dry reforming) utilizes two abundant greenhouse gases to produce industrially important synthesis gas (CO + H2 mixture). From this CO-H2 mixture, both fuels and valuable chemicals such as methanol can be produced using established technologies. The overall reaction proceeds through a series of surface intermediates:
CH4 + CO2 → 2CO + 2H2
Besides the mean route, methane direct decomposition, RWGS, methanation, Boudouard reaction (CO disproportionation to C and CO2) and carbon gasification can proceed, depending mainly on the bonding mode of the adsorbed CO formed. Methane activation primarily occurs on metal sites, whereas the nature of the intermediate formed is strongly influenced by the oxide support [32]. Consequently, species such as carboxylates, carbonates, formates, alkyls, and alkoxy intermediates may be present during methane reforming, depending on the catalyst structure (Table 3 and Table 4) [12,32,63,176]. Light alcohols, particularly methanol and ethanol, are also important feedstocks for hydrogen production via reforming reactions, with their behavior strongly dependent on the nature of the metal and the support [8,37,38,69,70,71,142,164,167,168,169,178,179,180,181]. One of the primary objectives in studying alcohol transformations is efficient hydrogen generation.
3.4.2. Methanol Steam Reforming
The decomposition and the steam reforming of methanol have been investigated on Pt-group metals supported by oxides, as well as on carbon-based supports such as Norit and multiwalled carbon. The main products are H2 and CO, with a very small amount of methane. The efficiency of the metals follows the order Ru > Rh > Ir ≥ Pt > Pd, while hydrogen selectivity above 623 K ranges between 60 and 95% [179].
CH3OH ↔ CO + 2H2
The dominant dissociation pathway of methanol involves the cleavage of the O–H bond, i.e., the formation of methoxy species [167,179,180], which is easily identified with different spectroscopies, including vibrational techniques (Table 4). Methoxy subsequently decomposes to CO and hydrogen. DFT studies indicate that formaldehyde can also form as an intermediate during methanol conversion [160]. In addition, XPS, SIMS, and TPD studies on Co, Ni, and Pd(100) surfaces suggest that partial C–O bond cleavage may occur, producing methyl species to some extent [72,168]. Important reactions are the following:
CH3OH(ads) → CH3O(ads) + H(ads)
CH3O(ads) → CO(ads) + 3H(ads)
CH3OH(ads) → CH3(ads) + OH(ads)
CH3O(ads) → (ads) + H2(g)
2CH3(ads) → 2C + 3H2(g)
CH3(ads) + H(ads) → CH4(g)
It was pointed out that the size-induced oxidation state of metals, including Pt, can influence the activity and selectivity of catalyst in methanol reactions. Under oxidizing conditions, formaldehyde is favored, especially when the Pt particle size is ~1 nm [15].
3.4.3. Ethanol Steam Reforming
Ethanol steam reforming has been widely studied on noble metal catalysts supported on oxides, particularly ceria-based systems, for sustainable hydrogen production [8,37,70,71,181]. Among these catalysts, Rh, Pt, and Ru exhibit the highest performance, with Rh showing superior activity due to its efficient C–C and C–H bond cleavage [6,33,62,157] and high resistance to carbon deposition [8]. The exceptional performance of Rh is further demonstrated in bimetallic systems, where the addition of a small amount of Rh (0.1%) to Co results in catalytic behavior, like for monometallic Rh supported on ceria [38]. In contrast, Pd/CeO2 and Au/CeO2 primarily produced acetaldehyde due to preferential C-H bond scission [162]. On oxide-free supports, such as Au on hexagonal boron nitride (h-BN) nanomesh supported on Rh(111), ethanol is converted to acetaldehyde with nearly 100% selectivity [163]. The selectivity of ethanol steam reforming is strongly influenced by the nature of the support [38]. Acidic supports such as Al2O3 favor ethylene formation, while SiO2 promotes acetaldehyde production. Ceria-based supports are most effective for hydrogen production. The ethanol conversion and H2 formation can be expressed by the following equation:
CH3CH2OH(ads) + 3H2O(ads) → 6H2(g) + 2CO2(g)
It is generally accepted that ethanol is activated on active sites of the catalysts, mainly on metallic components forming ethoxide. Its characteristic bands obtained by DRIFT can be seen in Table 4, together with their vibration modes, in agreement with several of the literature data. On noble metals (and some other metals) supported on ceria, the ethoxide can transform to acetaldehyde:
C2H5OH → C2H5O(ads) + H(ads)
C2H5O(ads) → CH3CHO(ads) + H(ads)
A certain part of acetaldehyde desorbs as the final product in the ethanol steam reforming; other fractions of acetaldehyde dehydrogenate form acetyl species where the C-O bond cleavage becomes more favorable. Although acetyl intermediates play an important role, their detection is challenging due to their short surface lifetime. Acetyl species were identified on Rh/CeO2 at 1684 cm−1 (C=O stretching mode) during ethanol transformation [170]. An easy C-C and C-H bond scissure has been confirmed earlier on Rh(111) and Pd(111) surfaces [69].
CH3CHO(ads) → CH3CO + H(ads)
CH3CO(ads) → CH3(ads) + CO(g)
2CH3(ads) → 2C(ads) + 3H2(gas)
Methane formation (Equation (81)) also occurs to a limited extent. Adsorbed acetaldehyde can undergo multiple reaction pathways: it can desorb, decompose to acyl, or react with hydroxyl groups or with lattice oxygen to form acetate. Acetate species are characterized by the νs(OCO) and νa(OCO) bands at 1452 and 1567 cm−1 [37,38,71,170,182]. Stable acetate surface product was also detected by near-ambient pressure XPS [93]. The findings show that acetate is a terminal and non-reactive product under the examined conditions. The acetate species were identical regardless of the supported metal (Pt, Pd, Ir, Rh, Ru), indicating that the formation of acetate species takes place on the oxide support [37,182]. Depending on metal and support and the nature of interface, the acetate may couple through aldol condensation to form acetone as byproduct at relatively high temperatures (from 720 K) [37,38].
CH3CHO(ads) + OH(ads) → CH3COO(ads) + H2
2CH3COO(ads) → CH3COCH3(g) + CO2(g) + O(ads)
Overall, spectroscopic and catalytic studies demonstrate that Pt-group metals, particularly Rh, are among the most effective catalysts for hydrogen production via ethanol reforming. However, the reaction pathway and efficiency are strongly influenced by oxide support. From the perspective of hydrogen production, ceria and ceria-modified supports appear to be the most favorable supports for noble metal catalysts [8,37,38,69,70,71,182].
4. Summary and Outlook
This review summarizes the identification and reactivity of intermediates formed in the CO + NO reaction, in the CO and CO2 hydrogenation and in the reforming of methane and light alcohols on metals and metal nanoparticles on oxide supports. The isocyanate surface complex is an existing intermediate in NO reduction with CO, and NCO is responsible for NH3 formation in real condition in exhaust gases. Alkyl groups and their reactions are important in the Fischer–Tropsch mechanism. CO2 hydrogenation proceeds via the formation and reaction of several intermediates which determine (together with the nature of active components of catalysts) the mechanism and the final products (CO, CH4, and CH3OH).
Some intermediates can be investigated during real catalytic conditions, but some intermediate surface complexes, due to their short lifetime, should be prepared from their parent’s molecules (HNCO, HCOOH, alkyl halogenates, alcohols, and aldehydes) via their surface dissociation (thermal- or photo-driven) at low temperature, where the intermediate species could be accumulated.
There is important information that not only the nature of metals, but the acidity, basicity and the reducibility of the supports basically influence the efficiency of the catalysts. In the future, investigations should be directed to the oxide-less support application (carbide, carbon, graphene, BN, BCN, Mo2S, etc.) or modified oxide-containing supports, where the disadvantageous metal–oxide interaction in the given reaction is hindered and a new route could be opened. Efforts to utilize RWGS reactions in growing carbon length to produce heavy hydrocarbons and oxygenated hydrocarbons are currently the subject of extensive research, with more planned in the immediate future [160,183]. Nowadays, a new strategy is being developed for planning scalable catalyst design, such as ZrO2-based catalysts for CO2 hydrogenation, emphasizing their multifunctional roles beyond conventional supports [184].
Currently, the M/CeO2 (M = Pt, Rh, Ir, Ru) seems to be the best catalyst for hydrogen production from ethanol reforming. Decreasing the formation of the adsorbed side-products (acetate), with suitable modification of support composition, could increase the selectivity to hydrogen production [71,182].
This review focuses mainly on thermal excitation. There is some evidence that photo-induced chemistry can open new reaction pathways, for example, in CO2 hydrogenation on Au/TiO2 catalysts [12]. In thermal reactions, the only product is CO. In UV photon-driven processes, the CH4 formation comes first, because the formate is also photon-sensitive and the activated formate can react with hydrogen, forming transiently to CH3O, which transforms to CH4 during further hydrogenation. Some heterosystems (for example, ZnO-FeOx) demonstrate high CO selectivity in CO2 reduction under visible and UV irradiation light [165]. DRIFTS confirmed formate and carbonyl species as key intermediates in the reaction mechanism. Photothermal catalysis, which couples photonic excitation with thermal driving forces, can alleviate the kinetic and thermodynamic barriers of CO2 activation and C-C coupling [166]. Photothermic-driven reactions have a bright future in technological applications. The results obtained by thermal-, photo- and electrochemical methods will increase the level of technology in this field in the near future.
The catalytic applications of single atoms, and nanoclusters below the nanosized range, give perspectives on the future directions toward developing a better understanding of the catalytic behavior of different metal entities for different types of reactions, including CO oxidation, selective oxidation–hydrogenation and photocatalytic reactions [9,10,11,12].
Author Contributions
J.K.: conceptualization, writing—review and editing, writing—original draft. I.S.: investigation, writing—original draft, formal analysis. A.E.: investigation, writing—original draft, formal analysis, validation. I.K.: investigation, writing—original draft, formal analysis. A.D.: visualization, validation. A.S.: conceptualization, project administration. Z.K.: resources, funding acquisition. All authors have read and agreed to the published version of the manuscript.
Funding
This research received no external funding.
Data Availability Statement
The original contributions presented in the study are included in the article, and further inquiries can be directed to the corresponding author.
Acknowledgments
The authors wish to thank the financial support of this work by the János Bolyai Research Scholarship Hungarian Academy of Sciences (I.S. BO/00369/25/7). This Project No. TKP2021-NVA-19 has been implemented with support from the Ministry of Innovation and Technology of Hungary from the National Research, Development, and Innovation Fund, financed under the TKP2021-NVA funding scheme. I.S. is grateful for EKÖP-24-4-SZTE-634, respectively, for the Research Fellowship Programme of Ministry of Culture and Innovation of Hungary. Project no. RRF-2.3.1-21-2022-00009, titled National Laboratory for Renewable Energy, has been implemented with the support provided by the Recovery and Resilience Facility of the European Union within the framework of Programme Széchenyi Plan Plus.
Conflicts of Interest
The authors declare no conflicts of interest.
Abbreviations
| TDS | Thermal desorption spectroscopy |
| TPD | Temperature-programmed desorption |
| AES | Auger electron spectroscopy |
| XPS | X-ray photoelectron spectroscopy |
| UPS | Ultraviolet photoelectron spectroscopy |
| IR | Infrared spectroscopy |
| FTIR | Fourier transform infrared spectroscopy |
| DRIFTS | Diffuse reflectance infrared Fourier transform spectroscopy |
| RAIRS | Reflection absorption infrared spectroscopy |
| SFG | Sum-frequency generation spectroscopy |
| HREELS | High-resolution electron energy-loss spectroscopy |
| EXAFS | Extended X-ray absorption fine structure |
| NAP-XPS | Near-ambient pressure X-ray photoelectron spectroscopy |
| SSITKA | Steady-state isotopic transient kinetic analysis |
| RWGS | Reverse water–gas shift (reaction) |
| DFT | Density functional theory |
| PE | Photoemission |
| EBF | Binding energy relative to the Fermi level |
| SMSI | Strong metal–support interaction |
References
- Ertl, G. Reactions on Solid Surfaces; Wiley: Hoboken, NJ, USA, 2010. [Google Scholar]
- Somorjai, G.A.; Li, Y. Introduction to Surface Chemistry and Catalysis; Wiley: Hoboken, NJ, USA, 2010. [Google Scholar]
- Van Santen, R.A. Modern Heterogeneous Catalysis: An Introduction; Wiley: Hoboken, NJ, USA, 2017. [Google Scholar]
- James, T.E.; Hemmingson, S.L.; Campbell, C.T. The energy of supported metal catalysts: From single atoms to large metal nanoparticles. ACS Catal. 2015, 5, 5673–5678. [Google Scholar] [CrossRef] [Scilit]
- Friend, C.M.; Xu, B. Heterogeneous Catalysis: A Central Science for a Sustainable Future. Acc. Chem. Res. 2017, 50, 517−521. [Google Scholar] [CrossRef] [Scilit]
- Zhao, K.; Wang, L.; Calizzi, M.; Moioli, E.; Züttel, A. In Situ Control of the Adsorption Species in CO2 Hydrogenation: Determination of Intermediates and Byproducts. J. Phys. Chem. C 2018, 122, 20888–20893. [Google Scholar] [CrossRef] [Scilit]
- Sun, S.; Sun, H.; Williams, P.T.; Wu, C. Recent advances in integrated CO2 capture and utilization: A review. Sustain. Energy Fuels 2021, 5, 4546–4559. [Google Scholar] [CrossRef] [Scilit]
- Silva, F.A.E.; do Marques, E.E.S.; Fonseca, F.C.; Gurlo, A.; Rodrigues, T.S. Noble metal-supported CeO2 catalysts for hydrogen production via ethanol steam reforming. ChemCatChem 2025, 17, e70002. [Google Scholar] [CrossRef] [Scilit]
- Li, Z.; Ji, S.; Liu, Y.; Cao, X.; Tian, S.; Chen, Y.; Niu, Z.; Li, Y. Well-defined materials for heterogeneous catalysis: From nanoparticles to isolated single-atom sites. Chem. Rev. 2020, 120, 623–682. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Sápi, A.; Rajkumar, T.; Kiss, J.; Kukovecz, Á.; Kónya, Z.; Somorjai, G.A. Metallic Nanoparticles in Heterogeneous Catalysis. Catal. Lett. 2021, 151, 2153–2175. [Google Scholar] [CrossRef] [Scilit]
- Liu, L.; Corma, A. Metal catalysts for heterogeneous catalysis from single atoms to nanoclusters and nanoparticles. Chem. Rev. 2018, 118, 4981–5079. [Google Scholar] [CrossRef] [Scilit]
- Kiss, J.; Kukovecz, Á.; Kónya, Z. Beyond nanoparticles: The role of sub-nanosized metal species in heterogeneous catalysis. Catal. Lett. 2019, 149, 1441–1454. [Google Scholar] [CrossRef] [Scilit]
- László, B.; Baán, K.; Zsuzsa Ferencz, Z.; Galbács, G.; Albert Oszkó, A.; Kónya, Z.; Kiss, J.; András Erdőhelyi, A. Gold Size Effect in the Thermal-Induced Reaction of CO2 and H2 on Titania- and Titanate Nanotube-Supported Gold Catalysts. J. Nanosci. Nanotechnol. 2019, 18, 470–477. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Bratlie, K.M.; Lee, H.; Komvopoulos, K.; Yang, P.; Somorjai, G.A. Platinum nanoparticle shape effects on benzene hydrogenation selectivity. Nano Lett. 2007, 7, 3097–3101. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Wang, H.; Wang, Y.; Zhu, Z.; Sapi, A.; An, K.; Kennedy, G.; Michalak, W.D.; Somorjai, G.A. Influence of Size-Induced Oxidation State of Platinum Nanoparticles on Selectivity and Activity in Catalytic Methanol Oxidation in the Gas Phase. Nano Lett. 2013, 13, 2976–2979. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Campbell, C.T. Ultrathin metal films and particles on oxide surfaces: Structural, electronic and chemisorptive properties. Surf. Sci. Rep. 1997, 27, 1–111. [Google Scholar] [CrossRef] [Scilit]
- Leybo, D.; Etim, U.J.; Monai, M.; Bare, S.R.; Zhong, Z.; Vogt, C. Metal–support interactions in metal-oxide-supported atomic, cluster and nanoparticle catalysis. Chem. Soc. Rev. 2024, 53, 10450–10490. [Google Scholar] [CrossRef] [Scilit]
- Gunter, P.L.J.; Niemantsverdriet, J.W.; Ribeiro, F.H.; Somorjai, G.A. Surface science approach to modeling supported catalysts. Catal. Rev. 1997, 39, 77–168. [Google Scholar] [CrossRef] [Scilit]
- Campbell, C.T. Catalyst-Support Interactions Electronic perturbations. Nat. Chem. 2012, 4, 597–598. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Pásztor, M. An infrared study of the influence of carbon monoxide chemisorption on the topology of supported rhodium. J. Phys. Chem. 1985, 89, 4789–4793. [Google Scholar] [CrossRef] [Scilit]
- Lu, Y.; Thompson, C.; Kuo, X.-Z.; Zhang, X.; Hoffman, A.S.; Boubnov, A.; Bara, S.R.; Kovarik, L.; Xin, H.; Karim, A.M. CO oxidation on MgAl2O4 supported Irn: Activation of lattice oxygen in the subnanometer regime and emergence of nuclearity-activity volcano. J. Mater. Chem. A 2022, 10, 4266–4278. [Google Scholar] [CrossRef] [Scilit]
- Berkó, A.; Szökő, J.; Solymosi, F. Effect of CO on the morphology of Pt nanoparticles supported on TiO2(1 1 0)-(1 × n). Surf. Sci. 2004, 566–568, 337–342. [Google Scholar] [CrossRef] [Scilit]
- Németh, R.; Kiss, J.; Solymosi, F. Surface Chemistry of HNCO and NCO on Pd(100). J. Phys. Chem. C 2007, 111, 1424–1427. [Google Scholar] [CrossRef] [Scilit]
- Kattel, S.; Liu, P.; Chen, J.G. Tuning Selectivity of CO2 Hydrogenation Reactions at the Metal/Oxide Interface. J. Am. Chem. Soc. 2017, 139, 9739−9754. [Google Scholar] [CrossRef] [Scilit]
- Freund, H.-J.; Roberts, M.W. Surface chemistry of carbon dioxide. Surf. Sci. Rep. 1996, 25, 225–273. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F. The bonding, structure and reactions of CO2 adsorbed on clean and promoted metal surfaces. J. Mol. Catal. 1991, 65, 337–358. [Google Scholar] [CrossRef] [Scilit]
- Taifan, W.; Boily, J.-F.; Baltrusaitis, J. Surface chemistry of carbon dioxide revisited. Surf. Sci. Rep. 2016, 71, 595–671. [Google Scholar] [CrossRef] [Scilit]
- Wang, X.; Shi, H.; Szanyi, J. Controlling selectivities in CO2 reduction through mechanistic understanding. Nat. Commun. 2017, 8, 513. [Google Scholar] [CrossRef] [Scilit]
- Guo, Y.; Mei, S.; Yuan, K.; Wang, D.J.; Liu, H.C.; Yan, C.H.; Zhang, Y.W. Low Temperature CO2 Methanation over CeO2-Supported Ru Single Atoms, Nanoclusters, and Nanoparticles Competitively Tuned by Strong Metal−Support Interactions and H-Spillover Effect. ACS Catal. 2018, 8, 6203–6215. [Google Scholar] [CrossRef] [Scilit]
- Szenti, I.; Efremova, A.; Kiss, J.; Sápi, A.; Óvári, L.; Halasi, G.; Haselmann, U.; Zhang, Z.; Morales-Vidal, J.; Baán, K.; et al. Pt/MnO Interface Induced Defects for High Reverse Water Gas Shift Activity. Angew. Chem. Int. Ed. 2024, 63, e202317343. [Google Scholar] [CrossRef] [Scilit]
- Ashcroft, A.T.; Cheetham, A.K.; Green, M.L.H.; Vernon, P.D.F. Partial oxidation of methane to synthesis gas using carbon dioxide. Nature 1991, 352, 225–226. [Google Scholar] [CrossRef] [Scilit]
- Erdőhelyi, A. Catalytic Reaction of Carbon Dioxide with Methane on Supported Noble Metal Catalysts. Catalysts 2021, 11, 159. [Google Scholar] [CrossRef] [Scilit]
- Fang, Y.; Wang, J.; Yao, K.; Sun, Y.; Hana, C.; Zhao, B. Regulating the content of Cu+ in Cu/CeO2 catalysts for the reverse water–gas shift reaction. New J. Chem. 2025, 49, 19337. [Google Scholar] [CrossRef] [Scilit]
- Huang, Z.; Li, Y.; Zhou, H.; Zeng, R.; Yang, Y.; Wang, L. Controlling CH4/CO Selectivity in CO2 Hydrogenation via Electronic Metal−Support Interaction at Ni-CeO2 Interfaces of Mg/Cu-Doped Catalysts. ACS Sustain. Chem. Eng. 2025, 13, 20070−20084. [Google Scholar] [CrossRef] [Scilit]
- Bent, B.E. Mimicking Aspects of Heterogeneous Catalysis: Generating, Isolating, and Reacting Proposed Surface Intermediates on Single Crystals in Vacuum. Chem. Rev. 1996, 96, 1361–1390. [Google Scholar] [CrossRef] [Scilit]
- Kovács, I.; Kiss, J.; Kónya, Z. The Potassium-Induced Decomposition Pathway of HCOOH on Rh(111). Catalysts 2020, 10, 675. [Google Scholar] [CrossRef] [Scilit]
- Mattos, L.V.; Jacobs, B.H.; Noronha, F.B. Production of Hydrogen from Ethanol: Review of Reaction Mechanism and Catalyst Deactivation. Chem. Rev. 2012, 112, 4094−4123. [Google Scholar] [CrossRef] [Scilit]
- Ferencz, Z.; Erdőhelyi, A.; Baán, K.; Oszkó, A.; Óvári, L.; Kónya, Z.; Papp, C.; Steinrück, H.-P.; Kiss, J. Effects of Support and Rh Additive on Co-Based Catalysts in the Ethanol Steam Reforming Reaction. ACS Catal. 2014, 4, 1205–1218. [Google Scholar] [CrossRef] [Scilit]
- Campbell, C.T. Energies of adsorbed catalytic intermediates on transition metal surfaces: Calorimetric measurements and benchmarks for theory. Acc. Chem. React. 2019, 52, 984–993. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Jamaati, M.; Torkashvand, M.; Sarabadani Tafreshi, S.; de Leeuw, N.H. A Review of Theoretical Studies on Carbon Monoxide Hydrogenation via Fischer-Tropsch Synthesis over Transition Metals. Molecules 2023, 28, 6525. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Nolen, M.A.; Tacey, S.A.; Know, S.; Farberow, C.A. Theoretical assessments of CO2 activation and hydrogenation pathways on transition-metal surfaces. Appl. Surf. Sci. 2023, 637, 157873. [Google Scholar] [CrossRef] [Scilit]
- Sun, C.; Sead, F.; Mirsalimova, S.; Natarajan, V.; Smerat, A.; Zhang, J.; Mahtab Alam, M.T.; Hengo, T. Mn, Co and Ti doped Nanocages (Mn2-C48, Co2-Si52 and Ti2-B30P30) and Mn, Co and Ti doped C-, Si- and BP-Nanotubes (5, 0), (6, 0) and (7, 0) as effective catalysts for CO2 reduction reaction. J. Phys. Chem. Solids 2026, 213, 113544. [Google Scholar] [CrossRef] [Scilit]
- Unland, M.L. Isocyanate intermediates in the Reaction NO + CO over a Pt/Al2O3 Catalyst. J. Phys. Chem. 1973, 77, 1952–1956. [Google Scholar] [CrossRef] [Scilit]
- Unland, M.L. Isocyanate Intermediates in Ammonia Formation over Noble Metal Catalysts for Automobile Exhaust Reactions. Science 1973, 179, 567–576. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Sárkány, J.; Schauer, A. Study of the formation of isocyanate surface complexes on Pt/Al2O3 catalysts. J. Catal. 1977, 46, 297–307. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Kiss, J.; Sárkány, J. On the Reaction of Surface Isocyanate over Platinum Catalyst. In Proceedings of the 3rd International Conference of Solid Surfaces, Vienna, Austria, 12–16 September 1977; pp. 819–822. [Google Scholar]
- Fischer, I.; Bell, A.T. A Comparative Study of CO and CO2 Hydrogenation over Rh/SiO2. J. Catal. 1996, 162, 54–65. [Google Scholar] [CrossRef] [Scilit]
- Kiss, J.; Solymosi, F. The Effect of Adsorbed Oxygen on the Stability of NCO on Rh(111) Studied by Reflection Absorption Infrared Spectroscopy. J. Catal. 1998, 179, 277–282. [Google Scholar] [CrossRef] [Scilit]
- Kiss, J.; Kis, A.; Solymosi, F. The Effects of Surface Additives on the Chemistry of CH3 on Rh(111) as Studied by Reflection Absorption Infrared Spectroscopy. Surf. Sci. 2000, 454–456, 273–279. [Google Scholar] [CrossRef] [Scilit]
- Zhou, X.L.; Liu, Z.-M.; Kiss, J.; Sloan, D.W.; White, J.M. Surface Chemistry of Chloroiodomethane, coadsorbed with H and O on Pt(111). J. Am. Chem. Soc. 1995, 117, 3565–3592. [Google Scholar] [CrossRef] [Scilit]
- Klivényi, G.; Solymosi, F. Generation of CH2 species: Thermal and photo-induced dissociation of CH2I2 on Rh(111) surface. Surf. Sci. 1995, 342, 168–184. [Google Scholar] [CrossRef] [Scilit]
- Bugyi, L.; Oszkó, A.; Solymosi, F. Oxidation of CH3 and C2H5 species on Rh(111). J. Catal. 1996, 159, 305–312. [Google Scholar] [CrossRef] [Scilit]
- Kiss, J.; Barthos, R.; Solymosi, F. The Effects of Potassium Overlayer on the Reaction Pathway of CH2 and C2H5 on Rh(111). Top. Catal. 2001, 14, 145–152. [Google Scholar] [CrossRef] [Scilit]
- Liu, Z.-M.; Zhou, X.-L.; Buchanan, D.A.; Kiss, J.; White, J.M. The Surface Chemistry of Vinyl Iodide. J. Am. Chem. Soc. 1992, 114, 2031–2039. [Google Scholar] [CrossRef] [Scilit]
- Kis, A.; Kiss, J.; Olasz, D.F.; Solymosi, F. Surface Reactions of Ethyl Groups on Clean and O-Modified Ru(001). J. Phys. Chem. B 2002, 106, 5221–5229. [Google Scholar] [CrossRef] [Scilit]
- Baltrusaitis, J.; Schuttlefield, J.; Zeitler, E.; Grassian, V.H. Carbon dioxide adsorption on oxide nanoparticle surfaces. Chem. Eng. J. 2011, 170, 471–481. [Google Scholar] [CrossRef] [Scilit]
- Kattel, S.; Yan, B.; Chen, J.G.C.; Liu, P. CO2 hydrogenation on Pt, Pt/SiO2 and Pt/TiO2: Importance of synergy between Pt and oxide support. J. Catal. 2016, 343, 115–126. [Google Scholar] [CrossRef] [Scilit]
- Vayssilov, G.N.; Mihaylov, M.; St. Petkov, P.; Konstantin, I.; Hadjiivanov, K.I.; Neyman, K.M. Reassignment of the Vibrational Spectra of Carbonates, Formates, and Related Surface Species on Ceria: A Combined Density Functional and Infrared Spectroscopy Investigation. J. Phys. Chem. C 2011, 115, 23435–23454. [Google Scholar] [CrossRef] [Scilit]
- Tóth, M.; Kiss, J.; Oszkó, A.; Pótári, G.; László, B.; Erdőhelyi, A. Hydrogenation of carbon dioxide on Au-Rh bimetallic clusters supported on titanate nanotubes, nanowires and TiO2. Top. Catal. 2012, 55, 747–756. [Google Scholar] [CrossRef] [Scilit]
- Wang, X.; Shi, H.; Kwak, J.H.; Szanyi, J. Mechanism of CO2 Hydrogenation on Pd/Al2O3 Catalysts: Kinetics and Transient DRIFTS-MS Studies. ACS Catal. 2015, 5, 6337–6349. [Google Scholar] [CrossRef] [Scilit]
- Wang, X.; Hong, Y.; Shi, H.; Szanyi, J. Kinetic modeling and transient DRIFTS–MS studies of CO2 methanation over Ru/Al2O3 catalysts. J. Catal. 2016, 343, 185–195. [Google Scholar] [CrossRef] [Scilit]
- Sápi, A.; Halasi, G.; Kiss, J.; Dobó, D.G.; Juhász, K.I.; Kolcsár, V.J.; Matolin, V.; Erdőhelyi, A.; Kukovecz, Á.; Kónya, Z. In-Situ DRIFTS and NAP-XPS Exploration of the Complexity of CO2 Hydrogenation Over Size Controlled Pt Nanoparticles Supported on Mesoporous NiO. J. Phys. Chem. C 2018, 122, 5553–5565. [Google Scholar] [CrossRef] [Scilit]
- László, B.; Baán, K.; Varga, E.; Oszkó, A.; Erdőhelyi, A.; Kónya, Z.; Kiss, J. Photo-induced reactions in the CO2-methane system on titanate nanotubes modified with Au and Rh nanoparticles. Appl. Catal. B Environ. 2016, 199, 473–484. [Google Scholar] [CrossRef] [Scilit]
- Falboa, L.; Viscontia, C.G.; Liettia, L.; Szanyi, J. The effect of CO on CO2 methanation over Ru/Al2O3 catalysts: A combined steady-state reactivity and transient DRIFT spectroscopy study. Appl. Catal. B Environ. 2019, 256, 117791. [Google Scholar] [CrossRef] [Scilit]
- Varga, G.; Sápi, A.; Varga, T.; Baán, K.; Szenti, I.; Halasi, G.; Mucsi, L.; Óvári, L.; Kiss, J.; Fogarassy, Z.; et al. Ambient pressure CO2 hydrogenation over a cobalt/manganese-oxide nanostructured interface: A combined in situ and ex situ study. J. Catal. 2020, 386, 70–80. [Google Scholar] [CrossRef] [Scilit]
- Efremova, A.; Rajkumar, T.; Szamosvölgyi, Á.; Sápi, A.; Baán, K.; Szenti, I.; Gómez-Pérez, J.F.; Varga, G.; Kiss, J.; Halasi, G.; et al. Complexity of a Co3O4 System under Ambient-Pressure CO2 Methanation: Influence of Bulk and Surface Properties on the Catalytic Performance. J. Phys. Chem. C 2021, 125, 7130–7141. [Google Scholar] [CrossRef] [Scilit]
- Efremova, A.; Szenti, I.; Kiss, J.; Szamosvölgyi, Á.; Sápi, A.; Baán, K.; Olivi, L.; Varga, G.; Fogarassy, Z.; Pécz, B.; et al. Nature of the Pt-Cobalt Oxide surface interaction and its role in the CO2 Methanation. Appl. Surf. Sci. 2022, 571, 15326. [Google Scholar] [CrossRef] [Scilit]
- Galhardo, T.S.; Braga, A.H.; Arpini, B.H.; Szanyi, J.; Gonçalves, R.V.; Zornio, B.F.; Miranda, C.R.; Rossi, L.M. Optimizing Active Sites for High CO Selectivity during CO2 Hydrogenation over Supported Nickel Catalysts. J. Am. Chem. Soc. 2021, 143, 4268–4280. [Google Scholar] [CrossRef] [Scilit]
- Houtman, C.J.; Barteau, M.A. Divergent Pathways of Acetaldehyde and Ethanol Decarbonylation on the Rh(111). Surf. J. Catal. 1991, 130, 528–546. [Google Scholar] [CrossRef] [Scilit]
- Raskó, J.M.; Dömök, M.; Baán, K.; Erdőhelyi, A. FTIR and mass spectrometric study of the interaction of ethanol and ethanol-water with oxide-supported platinum catalysts. Appl. Catal. A Gen. 2006, 299, 202–211. [Google Scholar] [CrossRef] [Scilit]
- Efremova, A.; Ballai, G.; Szamosvölgyi, Á.; Szenti, I.; Kutus, B.; Kiss, J.; Sápi, A.; Kukovecz, Á.; Kónya, Z. Interface Engineering of Pt–ZnO–CeO2 by Atomic Layer Deposition for Advanced Applications. Mater. Des. 2026, 263, 115640. [Google Scholar] [CrossRef] [Scilit]
- Steinbach, F.; Kiss, J.; Krall, R. Identification and Stability of CH3, CH2 and CH Species on Co and Ni Surfaces, a PES Investigation. Surf. Sci. 1985, 157, 401–412. [Google Scholar] [CrossRef] [Scilit]
- Zhou, X.-L.; Zhu, X.-Y.; White, J.M. Photochemistry at adsorbate/metal interfaces. Surf. Sci. Rep. 1991, 13, 73–220. [Google Scholar] [CrossRef] [Scilit]
- Jenks, C.J.; Bent, B.E. The Chemistry of Alkyl Iodides on Copper Surfaces. 1. Adsorption Geometry. J. Phys. Chem. B 2000, 104, 3008–3016. [Google Scholar] [CrossRef] [Scilit]
- Kovács, I.; Solymosi, F. Thermal and photoinduced dissociation of C2H5I to yield C2H5 on a Pd(100) Surface. J. Phys. Chem. 1993, 97, 11056–11063. [Google Scholar] [CrossRef] [Scilit]
- Kis, A.; Smith, K.C.; Kiss, J.; Solymosi, F. Adsorption and reactions of CH2I2 on Ru(001) surface. Surf. Sci. 2000, 460, 190–202. [Google Scholar] [CrossRef] [Scilit]
- Sexton, B.A.; Hughes, A.E.; Avery, N.R. A spectroscopic study of the adsorption and reactions of methanol, formaldehyde and methyl formate on clean and oxygenated Cu(110) surfaces. Surf. Sci. 1985, 155, 366–368. [Google Scholar] [CrossRef] [Scilit]
- Kovács, I.; Solymosi, F. Thermal and photoinduced dissociation of CH2I2 on Cu(100) surface. J. Phys. Chem. B 1997, 101, 5397–5404. [Google Scholar] [CrossRef] [Scilit]
- Kovács, I.; Solymosi, F. Thermal and photo-induced oxidation of CH2 on Cu (100). J. Mol. Catal. A Chem. 1999, 141, 31–38. [Google Scholar] [CrossRef] [Scilit]
- Jenks, C.J.; Bent, B.E.; Zaera, F. The Chemistry of Alkyl Iodides on Copper Surfaces. 2. Influence of Surface Structure on Reactivity. J. Phys. Chem. B 2000, 104, 3017–3027. [Google Scholar] [CrossRef] [Scilit]
- Kis, A.; Kiss, J.; Solymosi, F. Reaction of CH2 with Adsorbed O on Ru(001) Surface. Surf. Sci. 2000, 459, 149–160. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Kovács, I. Carbon-carbon coupling of methylene groups: Thermal and photo-induced dissociation of CH2I2 on Pd(100) surface. Surf. Sci. 1993, 296, 171–185. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Kiss, J. Interaction of HCOOH with Rhodium Surface Studied by Auger Electron, Electron Energy Loss and Thermal Desorption Spectroscopy. J. Catal. 1983, 81, 95–106. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Kovács, I. Adsorption and reaction of HCOOH on K-promoted Pd(100) surfaces. Surf. Sci. 1991, 259, 95–108. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Kovács, I.; Révész, K. Selective oxygen addition to adsorbed CH2 and CH3 on Pd(100). Surf. Sci. 1996, 356, 121–129. [Google Scholar] [CrossRef] [Scilit]
- Kiss, J.; Révész, K.; Solymosi, F. Photoelectron Spectroscopic Studies of the Adsorption of CO2 on Potassium-Promoted Rh(111) Surface. Surf. Sci. 1988, 207, 36–54. [Google Scholar] [CrossRef] [Scilit]
- Wohlrab, S.; Ehrlich, D.; Wambach, J.; Kuhlenbeck, H.; Freund, H.-J. Promoter action of alkali in the activation of CO2 on Pd (111): A HREELS case study. Surf. Sci. 1989, 220, 243–252. [Google Scholar] [CrossRef] [Scilit]
- Liu, Z.M.; Solymosi, F.; White, J.M. Spectroscopic study of K-induced activation of CO2 on Pt(111). Surf. Sci. 1991, 245, 289–304. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Klivényi, G. HREELS study on the formation of CO2− on K-promoted Rh(111) surfaces. Surf. Sci. 1994, 315, 255–268. [Google Scholar] [CrossRef] [Scilit]
- Papp, C.; Steinrück, H.-P. In situ high-resolution X-ray photoelectron spectroscopy –Fundamental insights in surface reactions. Surf. Sci. Rep. 2013, 68, 446–487. [Google Scholar] [CrossRef] [Scilit]
- Pantförder, P.; Zhu, J.Z.; Borgmann, D.; Denecke, R.; Steinrück, H.-P. New setup for in situ x-ray photoelectron spectroscopy from ultrahigh vacuum to 1 mbar. Rev. Sci. Inst. 2005, 76, 14102. [Google Scholar] [CrossRef] [Scilit]
- Varga, G.; Szenti, I.; Kiss, J.; Baán, K.; Halasi, G.; Óvári, L.; Szamosvölgyi, Á.; Mucsi, R.; Dodonyi, E.; Fogarassy, Z.; et al. Decisive role of Cu/Co interface in copper cobaltite derivates for high performance CO2 methanation. J. CO2 Util. 2023, 75, 102582. [Google Scholar] [CrossRef] [Scilit]
- Óvári, L.; Calderon, S.K.; Lykhach, Y.; Libuda, J.; Erdőhelyi, A.; Papp, C.; Kiss, J.; Steinrück, H.-P. Near ambient pressure XPS investigation of the interaction of ethanol with Co/CeO2(111). J. Catal. 2013, 307, 132–139. [Google Scholar] [CrossRef] [Scilit]
- Gong, X.-Q.; Liu, Z.-P.; Raval, R.; Hu, P. A Systematic Study of CO Oxidation on Metals and Metal Oxides: Density Functional Theory Calculations. J. Am. Chem. Soc. 2004, 126, 8–9. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Zhang, J.; Shu, M.; Niu, Y.; Yi, L.; Zhou, Y.; Zhao, S.; Tang, X.; Gao, F. Advances in CO catalytic oxidation on typical noble metal catalysts: Mechanism, performance and optimization. Chem. Eng. J. 2024, 495, 153523. [Google Scholar] [CrossRef] [Scilit]
- Langmuir, I. Part II.—“Heterogeneous reactions”. Chemical reactions on surfaces. Trans. Faraday Soc. 1922, 17, 607–620. [Google Scholar] [CrossRef] [Scilit]
- Prins, R. Eley-Riedel, the mechanism. Top. Catal. 2018, 61, 714–721. [Google Scholar] [CrossRef] [Scilit]
- Gao, F.; Wang, Y.; Goodman, D.W. CO/NO and CO/NO/O2 reactions over an Au–Pd single crystal catalyst. J. Catal. 2009, 268, 115–121. [Google Scholar] [CrossRef] [Scilit]
- Rainer, D.R.; Vesecky, S.M.; Koranne, M.; Oh, W.S.; Goodman, D.W. The CO + NO Reaction over Pd: A Combined Study Using Single-Crystal Planar-Model-Supported, and High-Surface-Area Pd/Al2O3 Catalysts. J. Catal. 1997, 167, 234–241. [Google Scholar] [CrossRef] [Scilit]
- Kyriakou, G.; Márquez, A.M.; Holgado, J.P.; Taylor, M.J.; Andrew, E.H.; Wheatley, A.E.H.; Mehta, J.P.; Sanz, J.F.; Simon, K.; Beaumont, S.K.; et al. Comprehensive Experimental and Theoretical Study of the CO + NO Reaction Catalyzed by Au/Ni Nanoparticles. ACS Catal. 2019, 9, 4919–4929. [Google Scholar] [CrossRef] [Scilit]
- Miners, J.H.; Bradshaw, A.M.; Gardner, P. Direct observation of surface isocyanate (NCO) formation during the CO + NO reaction on Pt{100}. Phys. Chem. Chem. Phys. 1999, 1, 4909–4912. [Google Scholar] [CrossRef] [Scilit]
- Hess, C.; Ozensoy, E.; Goodman, D.W. Combined in Situ and Infrared Kinetic Study of the Catalytic CO + NO Reaction on Pd(111) at Pressures up to 240 mbar. J. Phys. Chem. B 2003, 107, 2759. [Google Scholar] [CrossRef] [Scilit]
- Ozensoy, E.; Hess, C.; Goodman, D.W. Isocyanate formation in the catalytic reaction of CO+ NO on Pd (111): An in situ infrared spectroscopic study at elevated pressures. J. Am. Chem. Soc. 2002, 124, 8524–8525. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Ozensoy, E.; Goodman, D.W. Vibrational spectroscopic studies on CO adsorption, NO adsorption CO + NO reaction on Pd model catalysts. Phys. Chem. Chem. Phys. 2004, 6, 3765–3778. [Google Scholar] [CrossRef] [Scilit]
- Garda, G.R.; Castellani, N.J. Isocyanate (NCO) evidence in the CO + NO reaction over palladium. Appl. Catal. A Gen. 2015, 491, 48–56. [Google Scholar] [CrossRef] [Scilit]
- Sun, B.-Z.; Chen, W.-K.; Yi, Y.-J. Coadsorption of CO and NO on the Cu2O(111) surface: A periodic density functional theory study. J. Chem. Phys. 2009, 131, 174503. [Google Scholar] [CrossRef] [Scilit]
- Unland, M.I. Isocyanate intermediates in the reaction of NO and CO over noble metal catalysts. J. Catal. 1973, 31, 459–465. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Sárkány, J. An Infrared Study of the Surface Interaction between NO and CO on Rh/Al2O3 Catalysts. Appl. Surf. Sci. 1979, 3, 68–82. [Google Scholar] [CrossRef] [Scilit]
- Wangphon, C.; Khajondetchairit, P.; Rittiruam, M.; Ektarawong, A.; Alling, B.; Daengngern, R.; Praserthdam, P.; Saelee, T.; Praserthdam, S. Bridging the Gap in Understanding the Mechanism of NH3 Formation During NO Reduction by CO in the Presence of H2O Over Rh/Al2O3 Catalysts: DFT Study. Adv. Theory Simul. 2026, 9, e01922. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Völgyesi, L.; Sárkány, J. The effect of the support on the formation and stability of surface isocianate on platinum. J. Catal. 1978, 54, 336–344. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Kiss, J. Adsorption and decomposition of HNCO on Cu(111) surface studied by Auger electron, electron energy loss and thermal desorption spectroscopy. Surf. Sci. 1981, 104, 181–198. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Kiss, J. Adsorption and surface dissociation of HNCO on Pt(110) surfaces: LEED, AES, ELS and TDS studies. Surf. Sci. 1981, 108, 641–659. [Google Scholar] [CrossRef] [Scilit]
- Jones, J.E.; Trenary, M. Surface Chemistry of NCO Formed from HNCO on Pt(111). J. Phys. Chem. C 2008, 112, 20443–20450. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Berkó, A.; Tarnóczi, T.I. Effect of preadsorbed oxygen on the formation and decomposition of NCO on Rh(111) surfaces. Appl. Surf. Sci. 1984, 18, 233–245. [Google Scholar] [CrossRef] [Scilit]
- Farkas, A.P.; Berkó, A.; Solymosi, F. Interaction of HNCO with Au(111) surfaces. Surf. Sci. 2012, 606, 1345–1349. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Kiss, J. Interaction of C2N2 with Clean and Oxygen-Dosed Cu(111) Surface Studied by AES, ELS and TDS Measurements. Surf. Sci. 1981, 108, 368–380. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Bugyi, L. Adsorption and oxidation of C2N2 on oxygen dosed Rh(111) surfaces. Surf. Sci. 1984, 147, 685–701. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Berkó, A. Adsorption and oxidation of HCN on oxygen-dosed Cu(111) surface studied by AES, ELS and TDS measurements. Surf. Sci. 1982, 122, 275–291. [Google Scholar] [CrossRef] [Scilit]
- Gorte, R.J.; Schmidt, L.D.; Sexton, B.A. The electron energy loss spectrum of isocyanic acid on the Pt(111) Surface. J. Catal. 1981, 67, 387–391. [Google Scholar] [CrossRef] [Scilit]
- Dalla Betta, R.A.; Shelef, M. Isocyanates from the reaction of NO and CO on supported noble-metal catalysts. J. Mol. Catal. 1976, 1, 431–434. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Raskó, J. Infrared studies on the formation of isocyanate surface species over unsupported chromia. J. Catal. 1980, 65, 235–237. [Google Scholar] [CrossRef] [Scilit]
- Lorimer, D.; Bell, A.T. Reduction of NO by CO over a silica-supported platinum catalyst: Infrared and kinetic studies. J. Catal. 1979, 59, 223–238. [Google Scholar] [CrossRef] [Scilit]
- Biloen, P.; Sachtler, W.M.H. Mechanism of Hydrocarbon Synthesis over Fischer-Tropsch Catalysts. Adv. Catal. 1981, 30, 165–216. [Google Scholar] [CrossRef] [Scilit]
- Tauro, A.; Salomeone, F.; Clora, F.; Armandi, M.; Nodari, L.; Romagnoletti, L.; Felli, R.; Prone, R.; Bensaid, S. Effect of pre-treatment conditions on Fe-based catalyst for e-fuel production via modified Fischer-Tropsch synthesis. Chem. Eng. J. 2025, 51, 163154. [Google Scholar] [CrossRef] [Scilit]
- Zaera, F. Reversibity of C1 Hydrogenation-Dehydrogenation Reactions on Platinum Surfaces under Vacuum. Langmuir 1991, 7, 1998–1999. [Google Scholar] [CrossRef] [Scilit]
- Zaera, F. Molecular approach to the study of the mechanisms of alkyl reactions on metal surfaces. J. Mol. Catal. 1994, 86, 221–242. [Google Scholar] [CrossRef] [Scilit]
- Jeske, K.; Rösler, T.; Belleflamme, M.; Rodenas, T.; Fischer, N.; Claeys, M.; Leitner, W.; Vorholt, A.; Prieto, G. Direct Conversion of Syngas to Higher Alcohols via Tandem Integration of Fischer–Tropsch Synthesis and Reductive Hydroformylation. Angew. Chem. Int. Ed. 2022, 31, e202201004. [Google Scholar] [CrossRef] [Scilit]
- Mirzaei, A.A.; Sarani, R.H.; Azizi, R.S.; Vahid, S.; Torshizi, H.O. Kinetics modeling of Fischer-Tropsch synthesis on the unsupported Fe-Co-Ni (ternary) catalyst prepared using co-precipitation procedure. Fuel 2015, 140, 701–710. [Google Scholar] [CrossRef] [Scilit]
- van Santen, R.A.; Ciobica, I.M.; van Santen, E.; Ghouri, M.M. Chapter 3—Mechanistic Issues in Fischer–Tropsch Catalysis. Adv. Catal. 2011, 54, 127–187. [Google Scholar] [CrossRef] [Scilit]
- Lamont, C.L.A.; Conrad, H.; Bradshaw, A.M. The wavelength dependence of the photodissociation cross-section of CH3Br chemisorbed on Cu{111}. Surf. Sci. 1993, 280, 79–90. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Klivényi, G. HREELS study of CH3I and CH3 adsorbed on Rh(111) surface. J. Elect. Spect. Rel. Phenom. 1993, 64, 499–506. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Bugyi, L.; Oszkó, A. Generation of C2H5 species: Thermal and photo induced dissociation of C2H5I on Rh(111). Langmuir 1996, 12, 4145–4152. [Google Scholar] [CrossRef] [Scilit]
- Tjanda, S.; Zaera, F. The Effect of Hydrogen Coadsorption on the Thermal Chemistry of Methyl Iodide on Ni(100) Surfaces. J. Catal. 1993, 144, 361–376. [Google Scholar] [CrossRef] [Scilit]
- Scheer, K.C.; Kis, A.; Kiss, J.; White, J.M. Adsorption and reactions of CH2I2 on clean and oxygen-modified Ag(111): A RAIRS and TPD study. Top. Catal. 2002, 20, 43–51. [Google Scholar] [CrossRef] [Scilit]
- Kecskés, T.; Barthos, R.; Raskó, J.; Kiss, J. The effect of adsorbed CO on the surface chemistry of CH3 on Rh(111). Vacuum 2003, 71, 107–111. [Google Scholar] [CrossRef] [Scilit]
- Fairbrother, D.H.; Peng, X.D.; Trenary, M.; Stair, P.C. Surface Chemistry of Methyl Groups Adsorbed on Pt(111). J. Chem. Soc. Faraday Trans. 1995, 91, 3619−3625. [Google Scholar] [CrossRef] [Scilit]
- Yang, Q.Y.; Maynard, K.J.; Johnson, A.D.; Ceyer, S.T. The Structure and Chemistry of CH3 and CH Radicals Adsorbed on Ni(111). J. Chem. Phys. 1995, 102, 7734−7749. [Google Scholar] [CrossRef] [Scilit]
- Zhou, X.; Vejayan, H.; Beck, R.D.; Guo, H.; Jiang, B. Infrared Activities of Adsorbed Species on Metal Surfaces: The Puzzle of Adsorbed Methyl (CH3). J. Phys. Chem. Lett. 2021, 12, 11164–11169. [Google Scholar] [CrossRef] [Scilit]
- Hoffmann, H.; Griffiths, P.R.; Zaera, F. A RAIRS study on the surface chemistry of ethyl iodide on Pt(111). Surf. Sci. 1992, 262, 141–150. [Google Scholar] [CrossRef] [Scilit]
- Zaera, F.; Tjandra, S.; Janssen, T.V.W. Selectivity among Dehydrogenation Steps for Alkyl Groups on Metal Surfaces: Comparison between Nickel and Platinum. Langmuir 1998, 14, 1320–1327. [Google Scholar] [CrossRef] [Scilit]
- Barteau, M.A.; Broughton, J.Q.; Menzel, D. Vibrational spectroscopy of hydrocarbon intermediates on Ru(001). Appl. Surf. Sci. 1984, 19, 92–115. [Google Scholar] [CrossRef]
- Bol, C.W.J.; Friend, C.M. Controlling Selectivity in Alkyl Oxidation with Oxygen Coverage: The Reactions of Ethyl and 2-Propyl Iodide on Oxygen-Covered Rh(111). J. Phys. Chem. 1995, 99, 11930–11936. [Google Scholar] [CrossRef] [Scilit]
- Cui, G.; Lou, Y.; Hu, Y.; Zhou, M.; Li, Y.; Wang, Y.; Wang, W.; Li, J.; Jiang, G.; Xu, C. Enhancing CO2 Hydrogenation to Methanol at Tunable Cu−ZnO Interfaces on Hydrotalcite-Derived Cu Nanocatalysts. ACS Sustain. Chem. Eng. 2026, 14, 2445–2456. [Google Scholar] [CrossRef] [Scilit]
- Bol, C.W.J.; Friend, C.M.J. C-O Bond Formation by Direct Addition of Methyl Radicals to Surface Oxygen on Rh(111). Am. Chem. Soc. 1995, 117, 8053–8054. [Google Scholar] [CrossRef] [Scilit]
- Huberty, J.S.; Madix, R.J. An FTIR study of the bonding of methoxy on Ni(100): Effects of coadsorbed sulfur, carbon monoxide and hydrogen. Surf. Sci. 1996, 360, 144–156. [Google Scholar] [CrossRef] [Scilit]
- Anton, A.B.; Parmeter, J.E.; Weinberg, W.H. Adsorption of formaldehyde on the Ru(001) and Ru(001)-p(2 x2)O surfaces. J. Am. Chem. Soc. 1986, 108, 1823–1833. [Google Scholar] [CrossRef] [Scilit]
- Raskó, J.; Kecskés, T.; Kiss, J. Adsorption and reaction of formaldehyde on TiO2-supported Rh catalysts studied by FTIR and mass spectrometry. J. Catal. 2004, 226, 183–191. [Google Scholar] [CrossRef] [Scilit]
- Raskó, J.; Kiss, J. Adsorption and surface reactions of acetaldehyde on alumina-supported noble metal catalysts. Cat. Lett. 2005, 101, 71–77. [Google Scholar] [CrossRef] [Scilit]
- Kovács, I.; Ötvös, F.; Farkas, A.P.; Kiss, J.; Kónya, Z. A round dance of acetaldehyde molecular ensembles on Rh(111) surface; formation and decomposition of various paraldehyde conformers. J. Mol. Struct. 2022, 1264, 133311. [Google Scholar] [CrossRef] [Scilit]
- Zaera, F.; Guevremont, J.M.; Gleason, N.R.J. Direct Evidence for the Formation of a C−O Bond between Adsorbed Species. Phys. Chem. B 2001, 105, 2257–2259. [Google Scholar] [CrossRef] [Scilit]
- Boudart, M. Kinetics of Heterogeneous Catalysis Reactions; Princeton University Press: Princeton, NJ, USA, 1984. [Google Scholar]
- Van Etten, M.P.C.; De Laat, M.E.; Hensen, E.J.M.; Filot, I.A.W. Unraveling the Role of Metal−Support Interactions on the Structure Sensitivity of Fischer−Tropsch Synthesis. J. Phys. Chem. C 2023, 127, 15148–15156. [Google Scholar] [CrossRef] [Scilit]
- Dai, Y.; Zhu, C.; Wang, B.; Gao, M.; Ren, S.; Hou, Y.; Weize Wu, W. Carbon Dioxide Capture and In Situ Hydrogenation Conversion for the Production of C1 Chemicals. Carbon Hydrog. 2025, 27, 383–400. [Google Scholar] [CrossRef] [Scilit]
- Wang, W.; Wang, S.; Ma, X.; Gong, J. Recent advances in catalytic hydrogenation of carbon dioxide. Chem. Soc. Rev. 2011, 40, 3703–3727. [Google Scholar] [CrossRef] [Scilit]
- Hao, Z.; Li, Y.; Pan, Y.; Han, X.; Wang, Q.; Li, M.; Wang, Y.; Ma, X. Tuning the metal-support interaction via size-controlled ceria nanocubes in CO2 methanation over Ni/CeO2. Fuel 2026, 407, 137368. [Google Scholar] [CrossRef] [Scilit]
- Strekalova, A.; Shesterkina, A.A.; Beresnev, K.A.; Pribytkov, P.V.; Kapustin, G.I.; Mishin, I.V.; Kustov, L.M.; Kustov, A.L. Microwave Synthesis of Transition Metal (Fe, Co, Ni)-Supported Catalysts for CO2 Hydrogenation. Catalysts 2026, 16, 111. [Google Scholar] [CrossRef] [Scilit]
- Yang, J.; Gao, T.; Yang, J.; Xiao, F.; Liu, S.; Zhao, N.; Cao, D.; Li, H. Enhancement of Methanol Yield in CO2 Hydrogenation by Promoting CO2 Activation and Unlocking the RWGS + CO-Hydrogenation. ACS Catal. 2026, 16, 4773–4791. [Google Scholar] [CrossRef] [Scilit]
- Kostyniuk, A.; Yakushkin, S.; Likozar, B. Thermocatalytic CO2 hydrogenation to high-yield ethanol and C3–C5 alcohols over promoted Fe-based catalysts. J. Energy Chem. 2026, 116, 322–338. [Google Scholar] [CrossRef] [Scilit]
- Ma, J.; Sun, N.; Zhang, X.; Zhao, N.; Xiao, F.; Wei, W.; Sun, Y. A short review of catalysis for CO2 conversion. Cat. Today 2009, 148, 221–231. [Google Scholar] [CrossRef] [Scilit]
- Polinskí, M.P.; Kozlov, S.M. Reaction network of CO2 hydrogenation into C1–2 oxygenates and its BEP relationships. EES Catal. R. Chem. 2026. [Google Scholar] [CrossRef] [Scilit]
- Feng, L.; Zhao, J.-W.; Wei, W.-Y.; Wang, H.-Y.; Jiang, Y.-Q.; Liu, J.-X.; Li, W.-X. Nature of Reverse Water−Gas Shift Reactions at Metal−Oxide Interfaces Uncovered via Interpretable Machine Learning. J. Am. Chem. Soc. 2026, 148, 3602−3613. [Google Scholar] [CrossRef] [Scilit]
- Kiss, J.; Sápi, A.; Tóth, M.; Kukovecz, Á.; Kónya, Z. Rh-Induced Support Transformation and Rh Incorporation in Titanate Structures and Their Influence on Catalytic Activity. Catalysts 2020, 10, 212. [Google Scholar] [CrossRef] [Scilit]
- He, Y.; Müller, F.H.; Palkovits, R.; Zeng, F.; Mebrahtu, C. Tandem catalysis for CO2 conversion to higher alcohols: A review. Appl. Catal. B Environ. Energy 2024, 345, 123663. [Google Scholar] [CrossRef] [Scilit]
- László, B.; Baán, K.; Oszkó, A.; Erdőhelyi, A.; Kiss, J.; Kónya, Z. Hydrogen evolution in the photocatalytic reaction between methane and water in the presence of CO2 on titanate and titania supported Rh and Au catalysts. Top. Catal. 2018, 61, 875–888. [Google Scholar] [CrossRef] [Scilit]
- Basheer, H.S.; Yadav, M.; Szamosvölgyi, Á.; Balog, Á.; Janáky, C.; Efremova, A.; Kiss, J.; András Sápi, A.; Ákos Kukovecz, Á.; Kónya, Z. Interface Engineering of Fe-Loaded ZnO Nanostructures via Wet Impregnation and Hydrothermal Routes for Enhanced CO Selectivity in CO2 Photoreduction. ChemSusChem 2026, 19, 202502004. [Google Scholar] [CrossRef] [Scilit]
- Bafaqeer, A.; Elhussien, A.A.; Almansaf, Z.; Jaseer, E.A.; Amin, N.A.S.; Fouad, W.A. Photothermal catalysis for CO2 hydrogenation to C2+ products: Mechanisms, catalytic system, and selectivity control. J. Environ. Chem. Eng. 2026, 14, 121154. [Google Scholar] [CrossRef] [Scilit]
- Greeley, J.; Mavrikakis, M. Competitive Paths for Methanol Decomposition on Pt(111). J. Am. Chem. Soc. 2004, 126, 3910–3919. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Guo, X.; Hanley, L.; Yates, J.T., Jr. Thermal stability of the carbon-oxygen bond of methanol on the palladium(111) surface: An isotopic mixing study. J. Am. Chem. Soc. 1989, 111, 3155–3157. [Google Scholar] [CrossRef] [Scilit]
- Farkas, A.P.; Szitás, Á.; Jurdi, D.; Palotás, K.; Kiss, J.; Kónya, Z. Selective Transformation of Ethanol to Acetaldehyde Catalyzed by Au/h-BN Interface Prepared on Rh(111) Surface. Appl. Catal. A Gen. 2020, 502, 117440. [Google Scholar] [CrossRef] [Scilit]
- Tóth, M.; Varga, E.; Oszkó, A.; Baán, K.; Kiss, J.; Erdőhelyi, A. Partial oxidation of ethanol on supported Rh catalysts: Effect of the oxide support. J. Mol. Catal. A Chem. 2016, 411, 377–387. [Google Scholar] [CrossRef] [Scilit]
- Óvári, L.; Farakas, A.P.; Palotás, K.; Vári, G.; Szenti, I.; Berkó, A.; Kiss, J.; Kónya, Z. Hexagonal boron nitride on metal surfaces as a support and template. Surf. Sci. Rep. 2024, 79, 100637. [Google Scholar] [CrossRef] [Scilit]
- Hegedűs, T.; Szenti, I.; Efremova, A.; Szamosvölgyi, Á.; Baán, K.; Kiss, J.; Kónya, Z. Hexagonal boron nitride fibers as ideal catalytic support to experimentally measure the distinct activity of Pt nanoparticles in CO2 hydrogenation. Heliyon 2024, 10, e40078. [Google Scholar] [CrossRef] [Scilit]
- Cao, A.; Wang, Z.; Li, H.; Elnabawy, A.O.; Nørskov, J.K. New insights on CO and CO2 hydrogenation for methanol synthesis: The key role of adsorbate-adsorbate interactions on Cu and the highly active MgO-Cu interface. J. Catal. 2021, 400, 325–331. [Google Scholar] [CrossRef] [Scilit]
- Solymosi, F.; Kiss, J. Adsorption of H2O on Clean and Boron-Contaminated Rh Surfaces. Surf. Sci. 1986, 177, 191–206. [Google Scholar] [CrossRef] [Scilit]
- Szamosvölgyi, Á.; Rajkumar, T.; Sápi, A.; Szenti, I.; Ábel, M.; Gomez-Perez, J.F.; Baán, K.; Fogarassy, Z.; Dodonyi, E.; Kiss, J.; et al. Interfacial Ni active sites strike solid solutional counterpart in CO2 hydrogenation. Environ. Tech. Innov. 2022, 27, 102747. [Google Scholar] [CrossRef] [Scilit]
- Bartholomew, C.H. Mechanisms of catalyst deactivation. Appl. Catal. A Gen. 2001, 212, 17–60. [Google Scholar] [CrossRef] [Scilit]
- Liu, W.; Zhao, H.; Wu, X.; Wu, J.; Chou, L.; Dury, G.; Hu, W.; Polynski, M.V.; Subramanian, A.; Kozlov, S.M.; et al. Selectivity Anomaly in CO2 Hydrogenation over In−Pd Intermetallic Compounds. ACS Catal. 2026, 16, 3464–3478. [Google Scholar] [CrossRef] [Scilit]
- Lai, W.; Wang, L.; Dai, Z.; Yao, L.; Jiang, W. The mitigation of carbon deposition for Ni-based catalyst in CO2 reforming of methane: A combined experimental and DFT study. Carbon Cap. Sci. Technol. 2024, 13, 100286. [Google Scholar] [CrossRef] [Scilit]
- Tolmacsov, P.; Gazsi, A.; Solymosi, F. Decomposition and reforming of methanol on Pt metals supported by carbon Norit. Appl. Catal. A Gen. 2009, 362, 58–63. [Google Scholar] [CrossRef] [Scilit]
- Badmaev, S.D.; Belyaev, V.D.; Potemkin, D.I.; Snytnikov, P.V.; Sobyanin, V.A.; Kharton, V.V. Methanol Decomposition to Synthesis Gas over Supported Platinum-Containing Catalysts. Catal. Ind. 2023, 15, 367–373. [Google Scholar] [CrossRef] [Scilit]
- Yee, A.S.; Morrison, S.J.; Idriss, H. The reactions of ethanol over M/CeO2 catalysts: Evidence of carbon–carbon bond dissociation at low temperatures over Rh/CeO2. Catal. Today 2000, 63, 327–335. [Google Scholar] [CrossRef] [Scilit]
- Erdőhelyi, A.; Raskó, J.; Kecskés, T.; Tóth, M.; Dömök, M.; Baán, K. Hydrogen formation in ethanol reforming on supported noble metal catalysts. Catal. Today 2006, 116, 367–376. [Google Scholar] [CrossRef] [Scilit]
- Najari, S.; Saeidi, S.; Sápi, A.; Kónya, Z.; Somorjai, G.A. Unveiling the Power of Proximity of Prevalent Fe-Based Tandem Catalysts in CO2 Hydrogenation via Modified Fischer−Tropsch: Crucial Relations toward Industrialization. Chem. Rev. 2025, 125, 10179–10247. [Google Scholar] [CrossRef] [Scilit]
- Deng, H.; Luo, F.; Feng, G.; Zhang, R.; Ye, R. Research Progress and Prospects of ZrO2-Based Catalysts in CO2 Hydrogenation. Carbon Hydrog. 2025, 27, 401–419. [Google Scholar] [CrossRef] [Scilit]
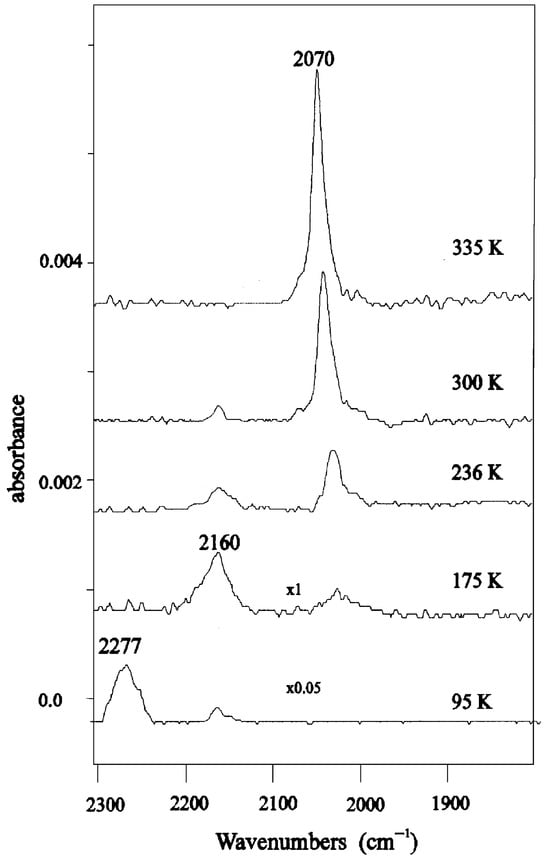
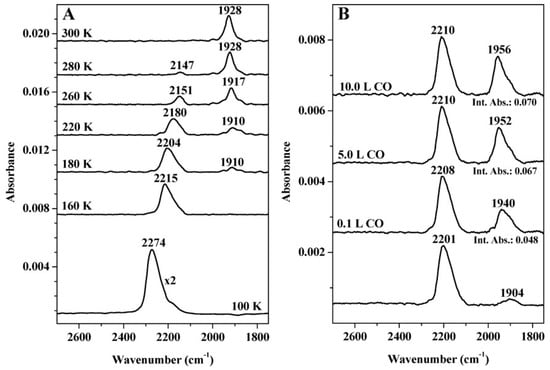
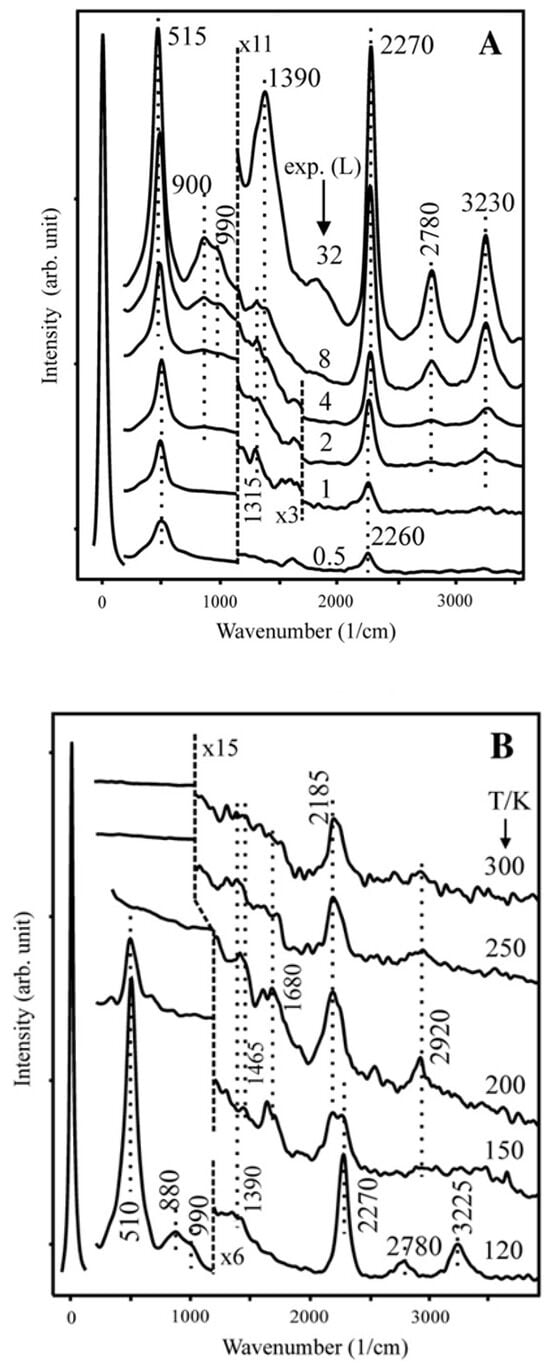
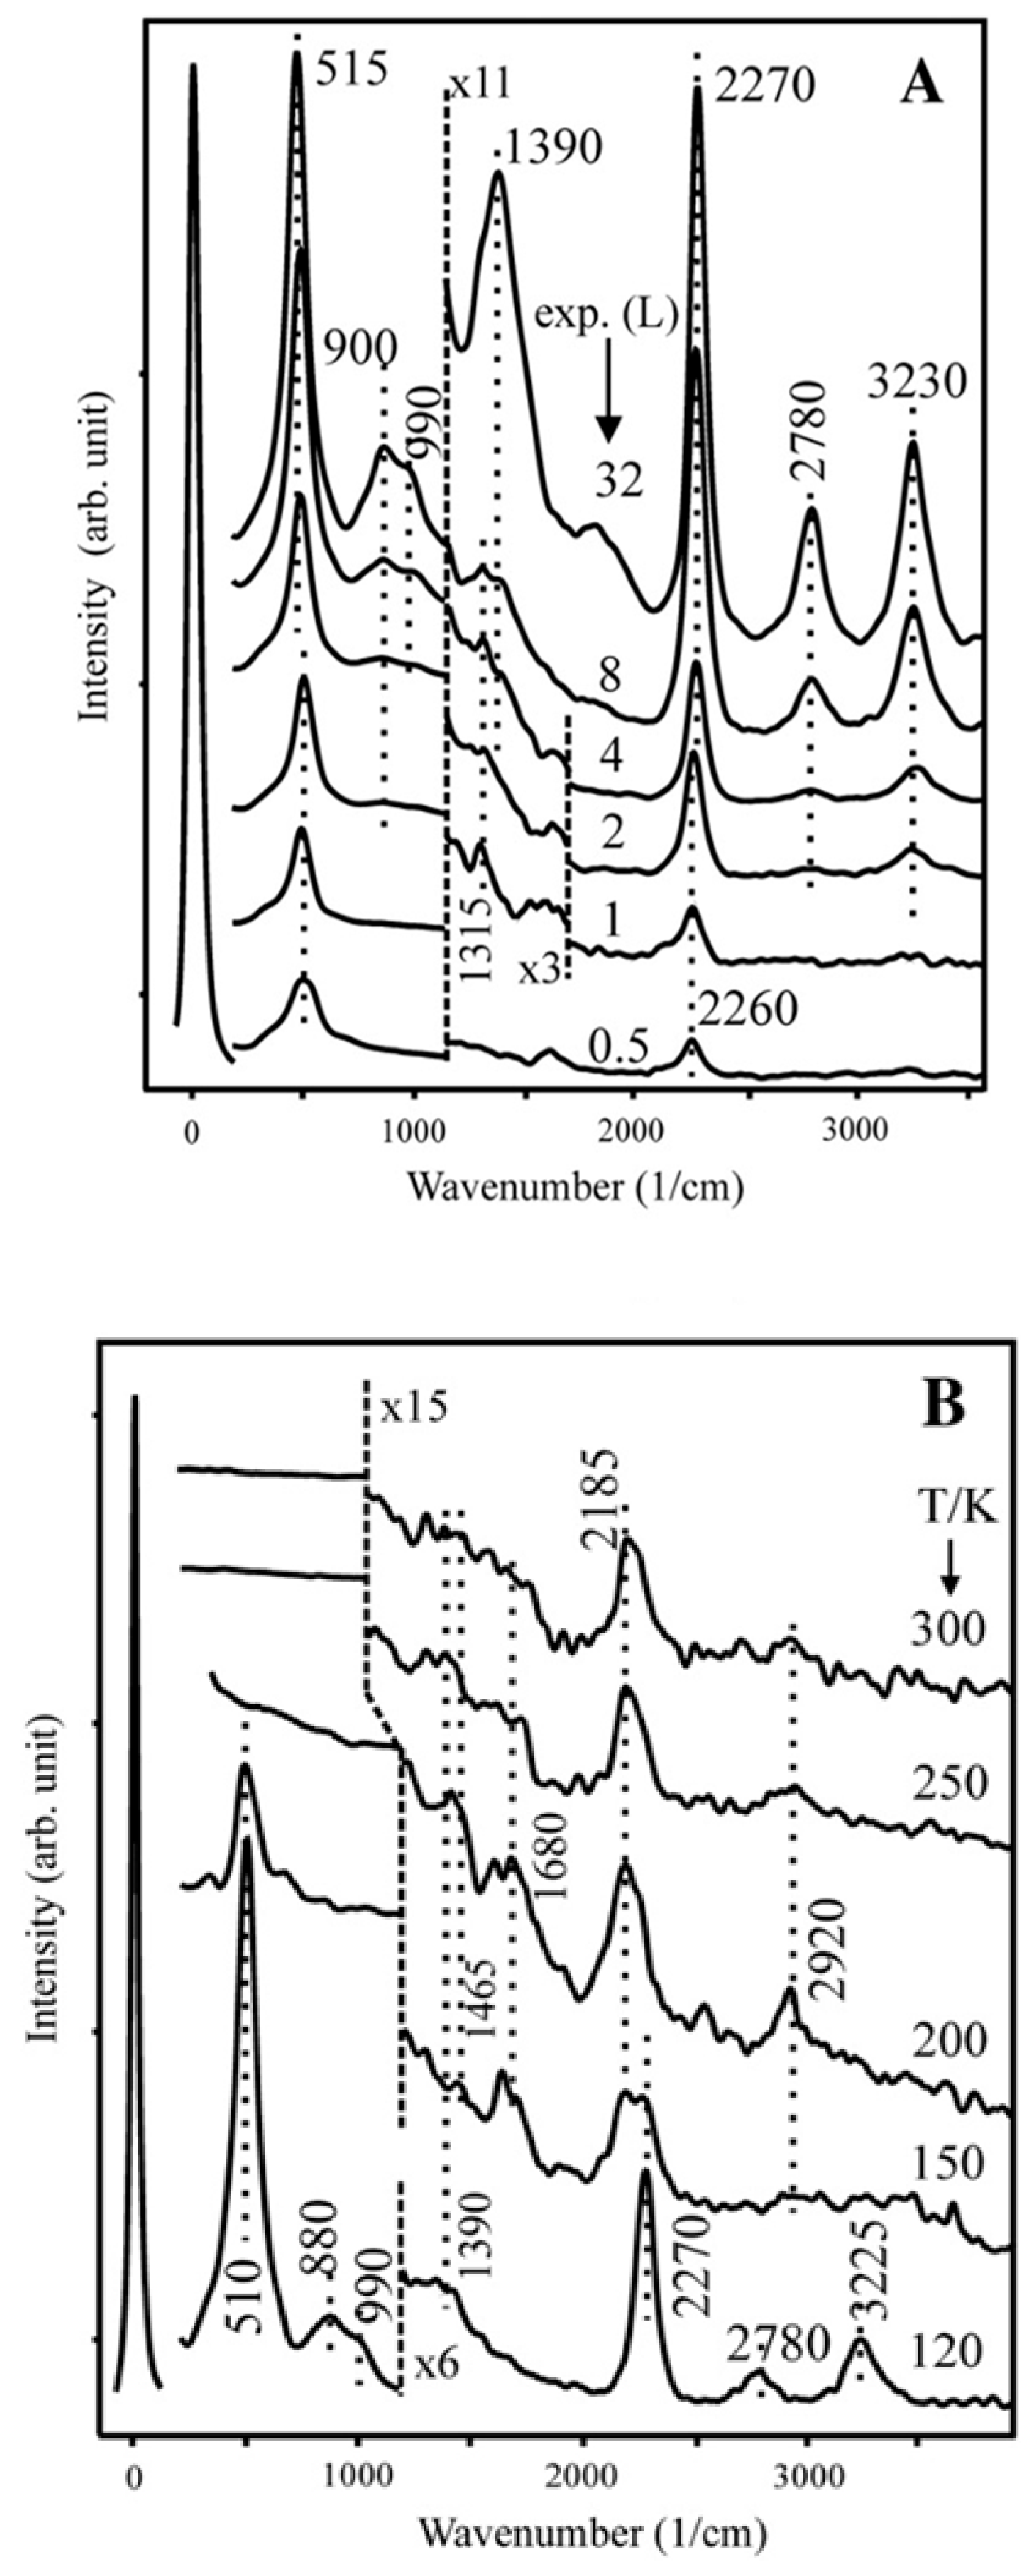
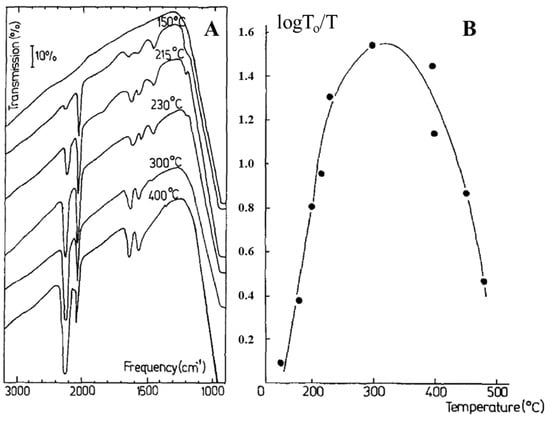
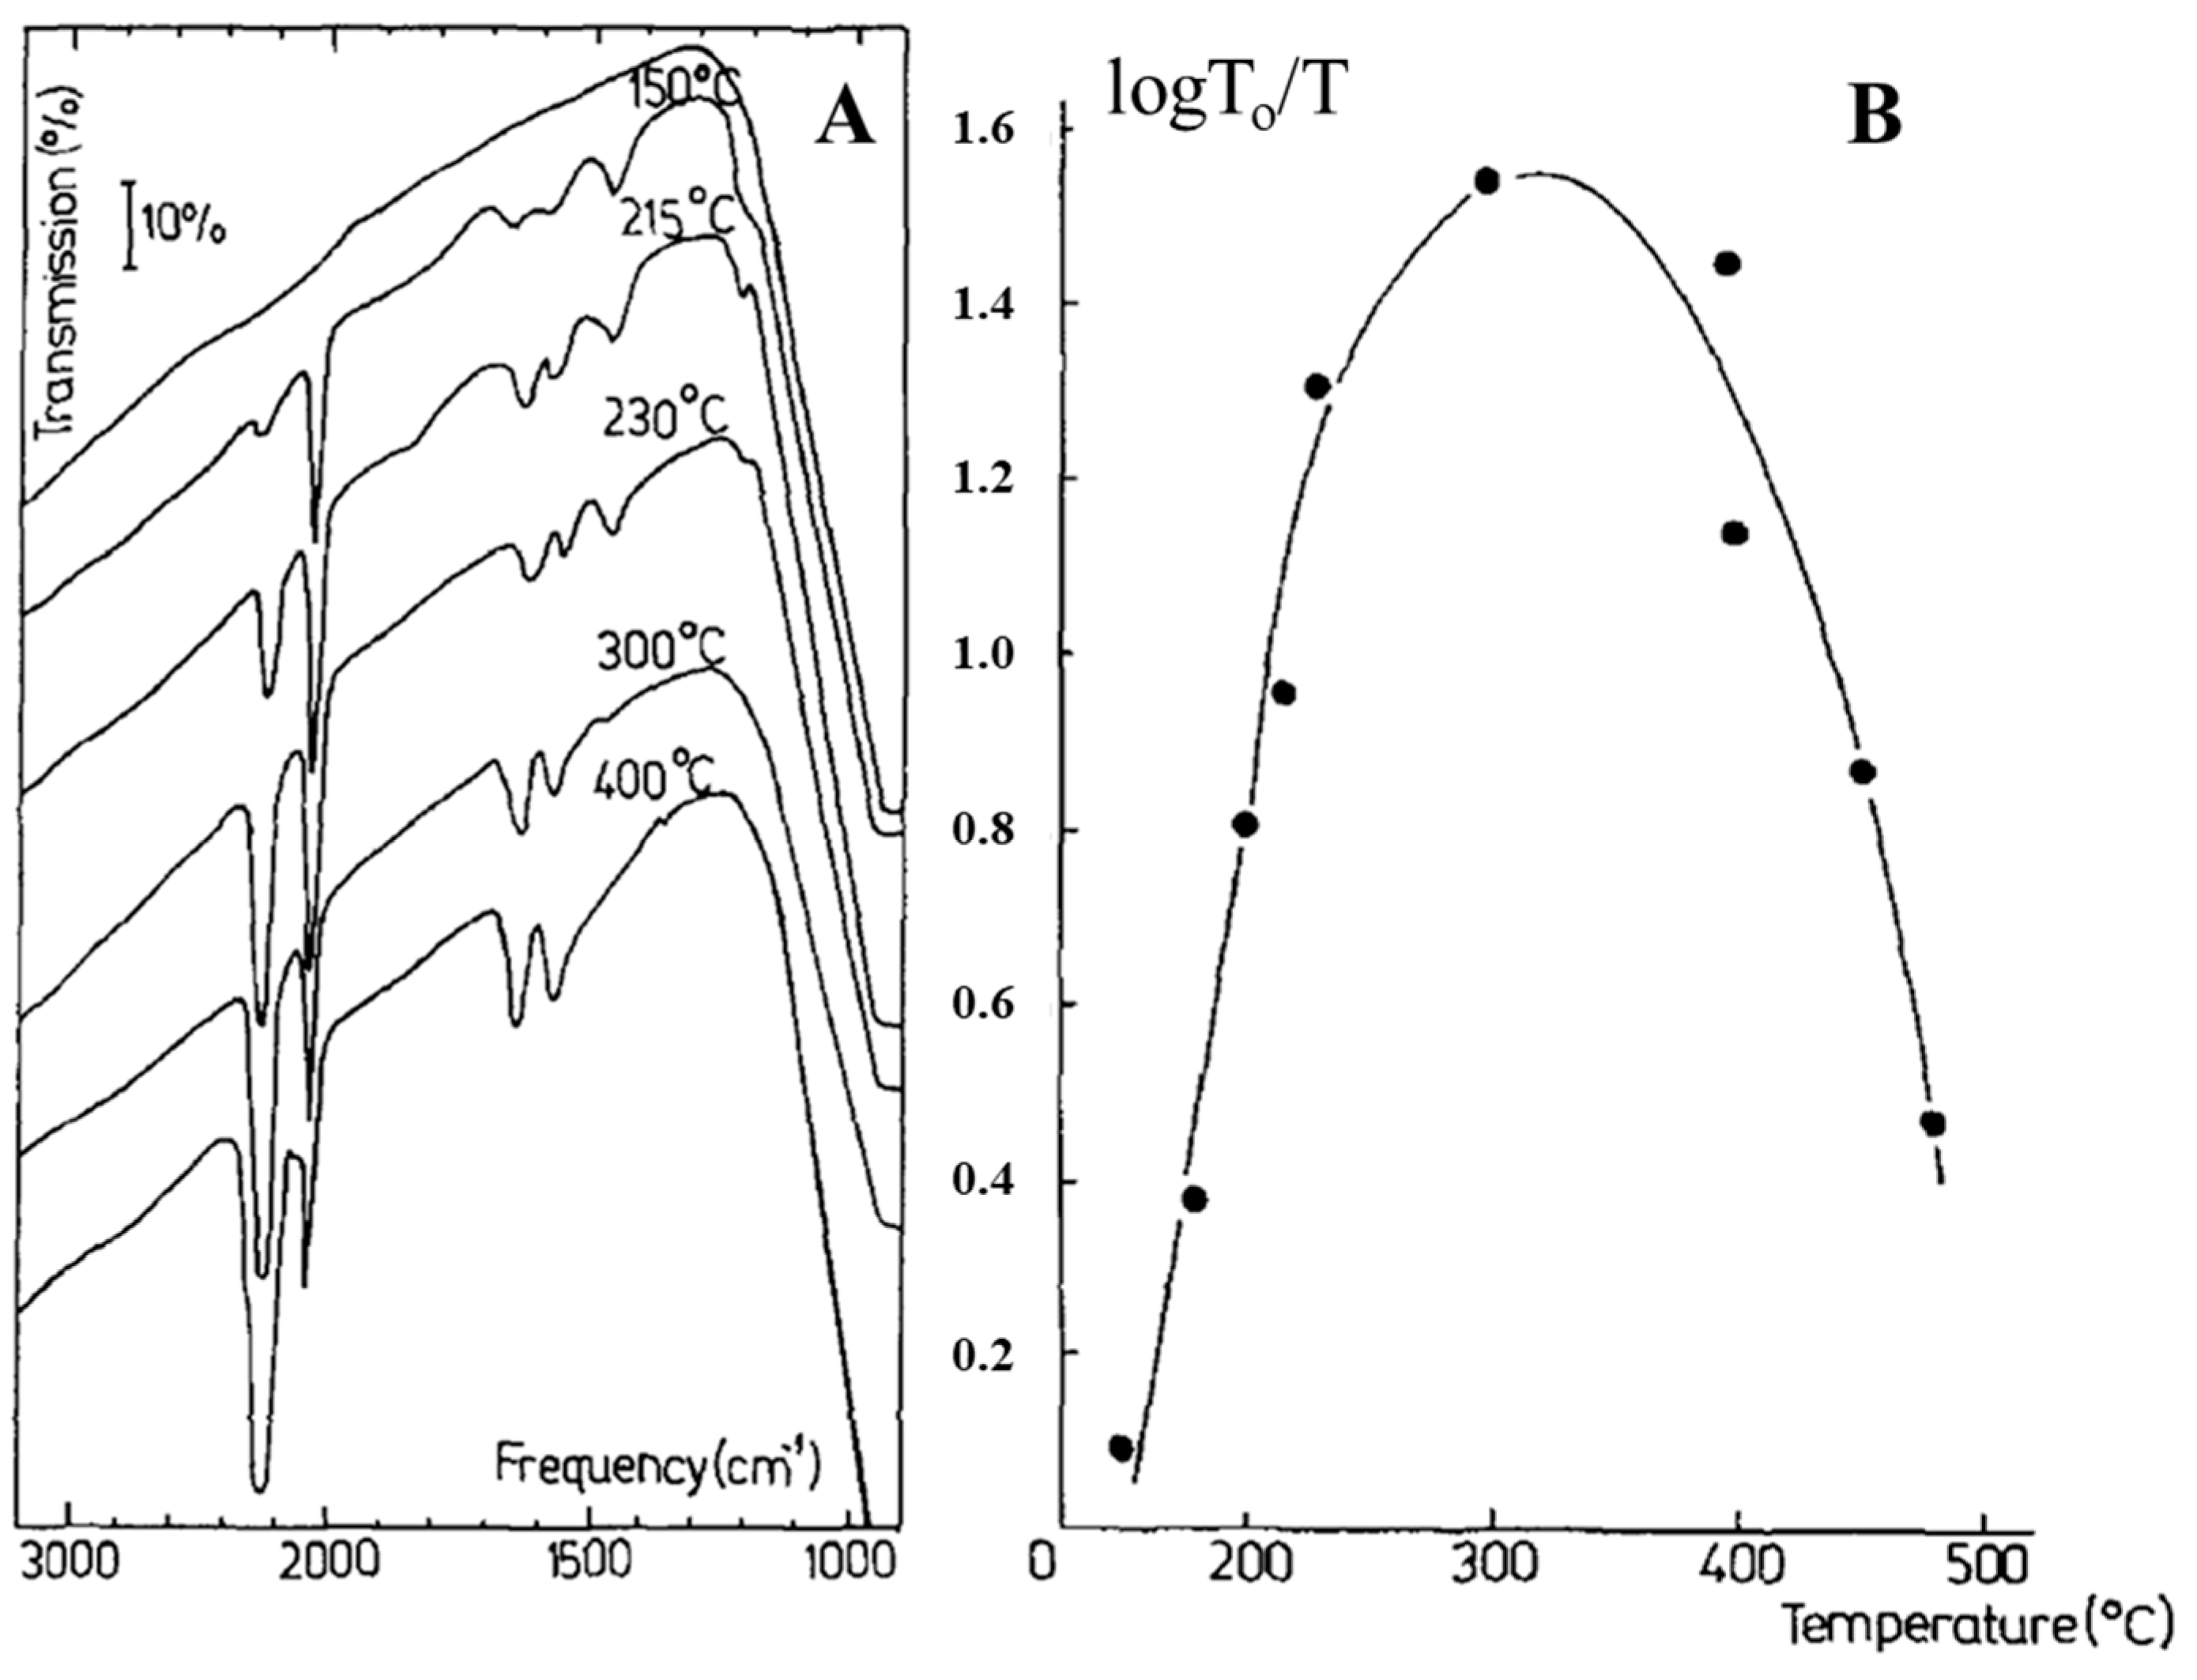
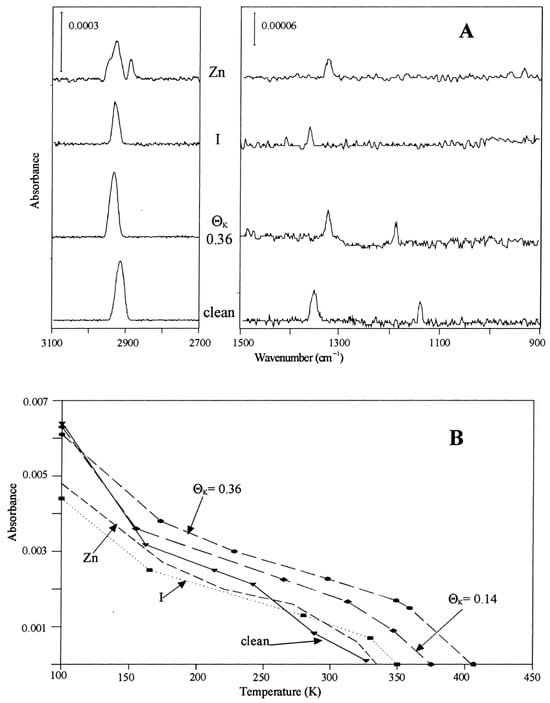
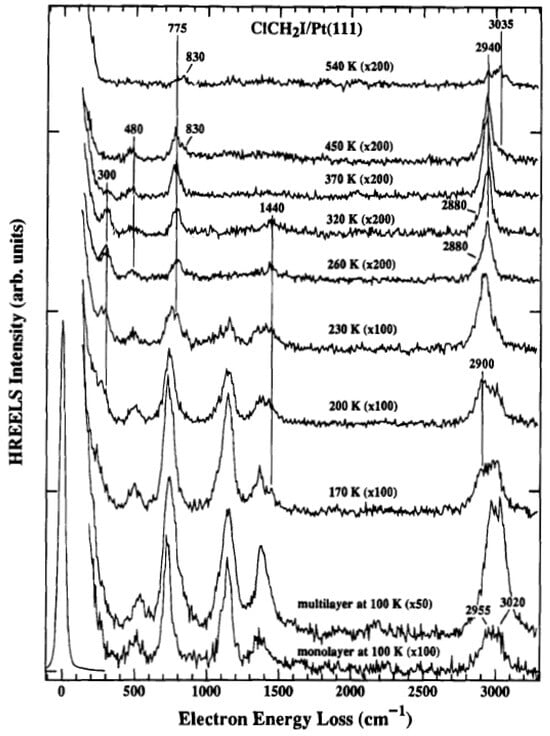
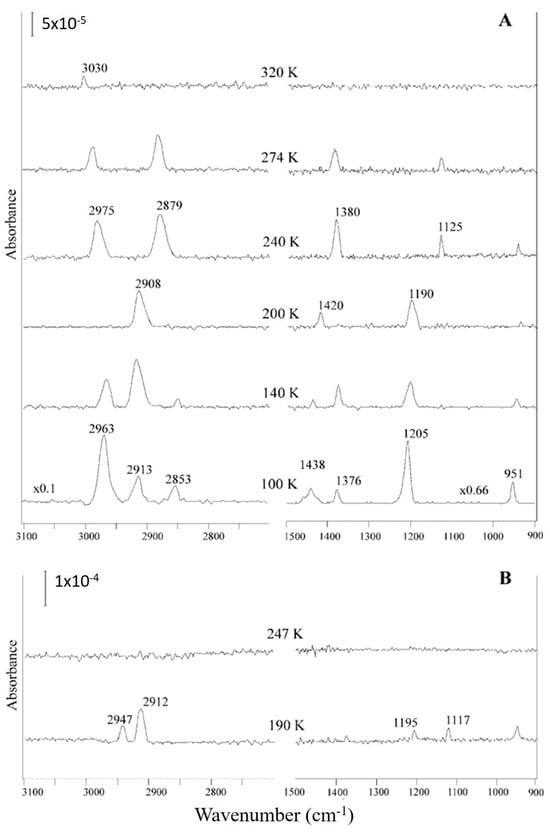
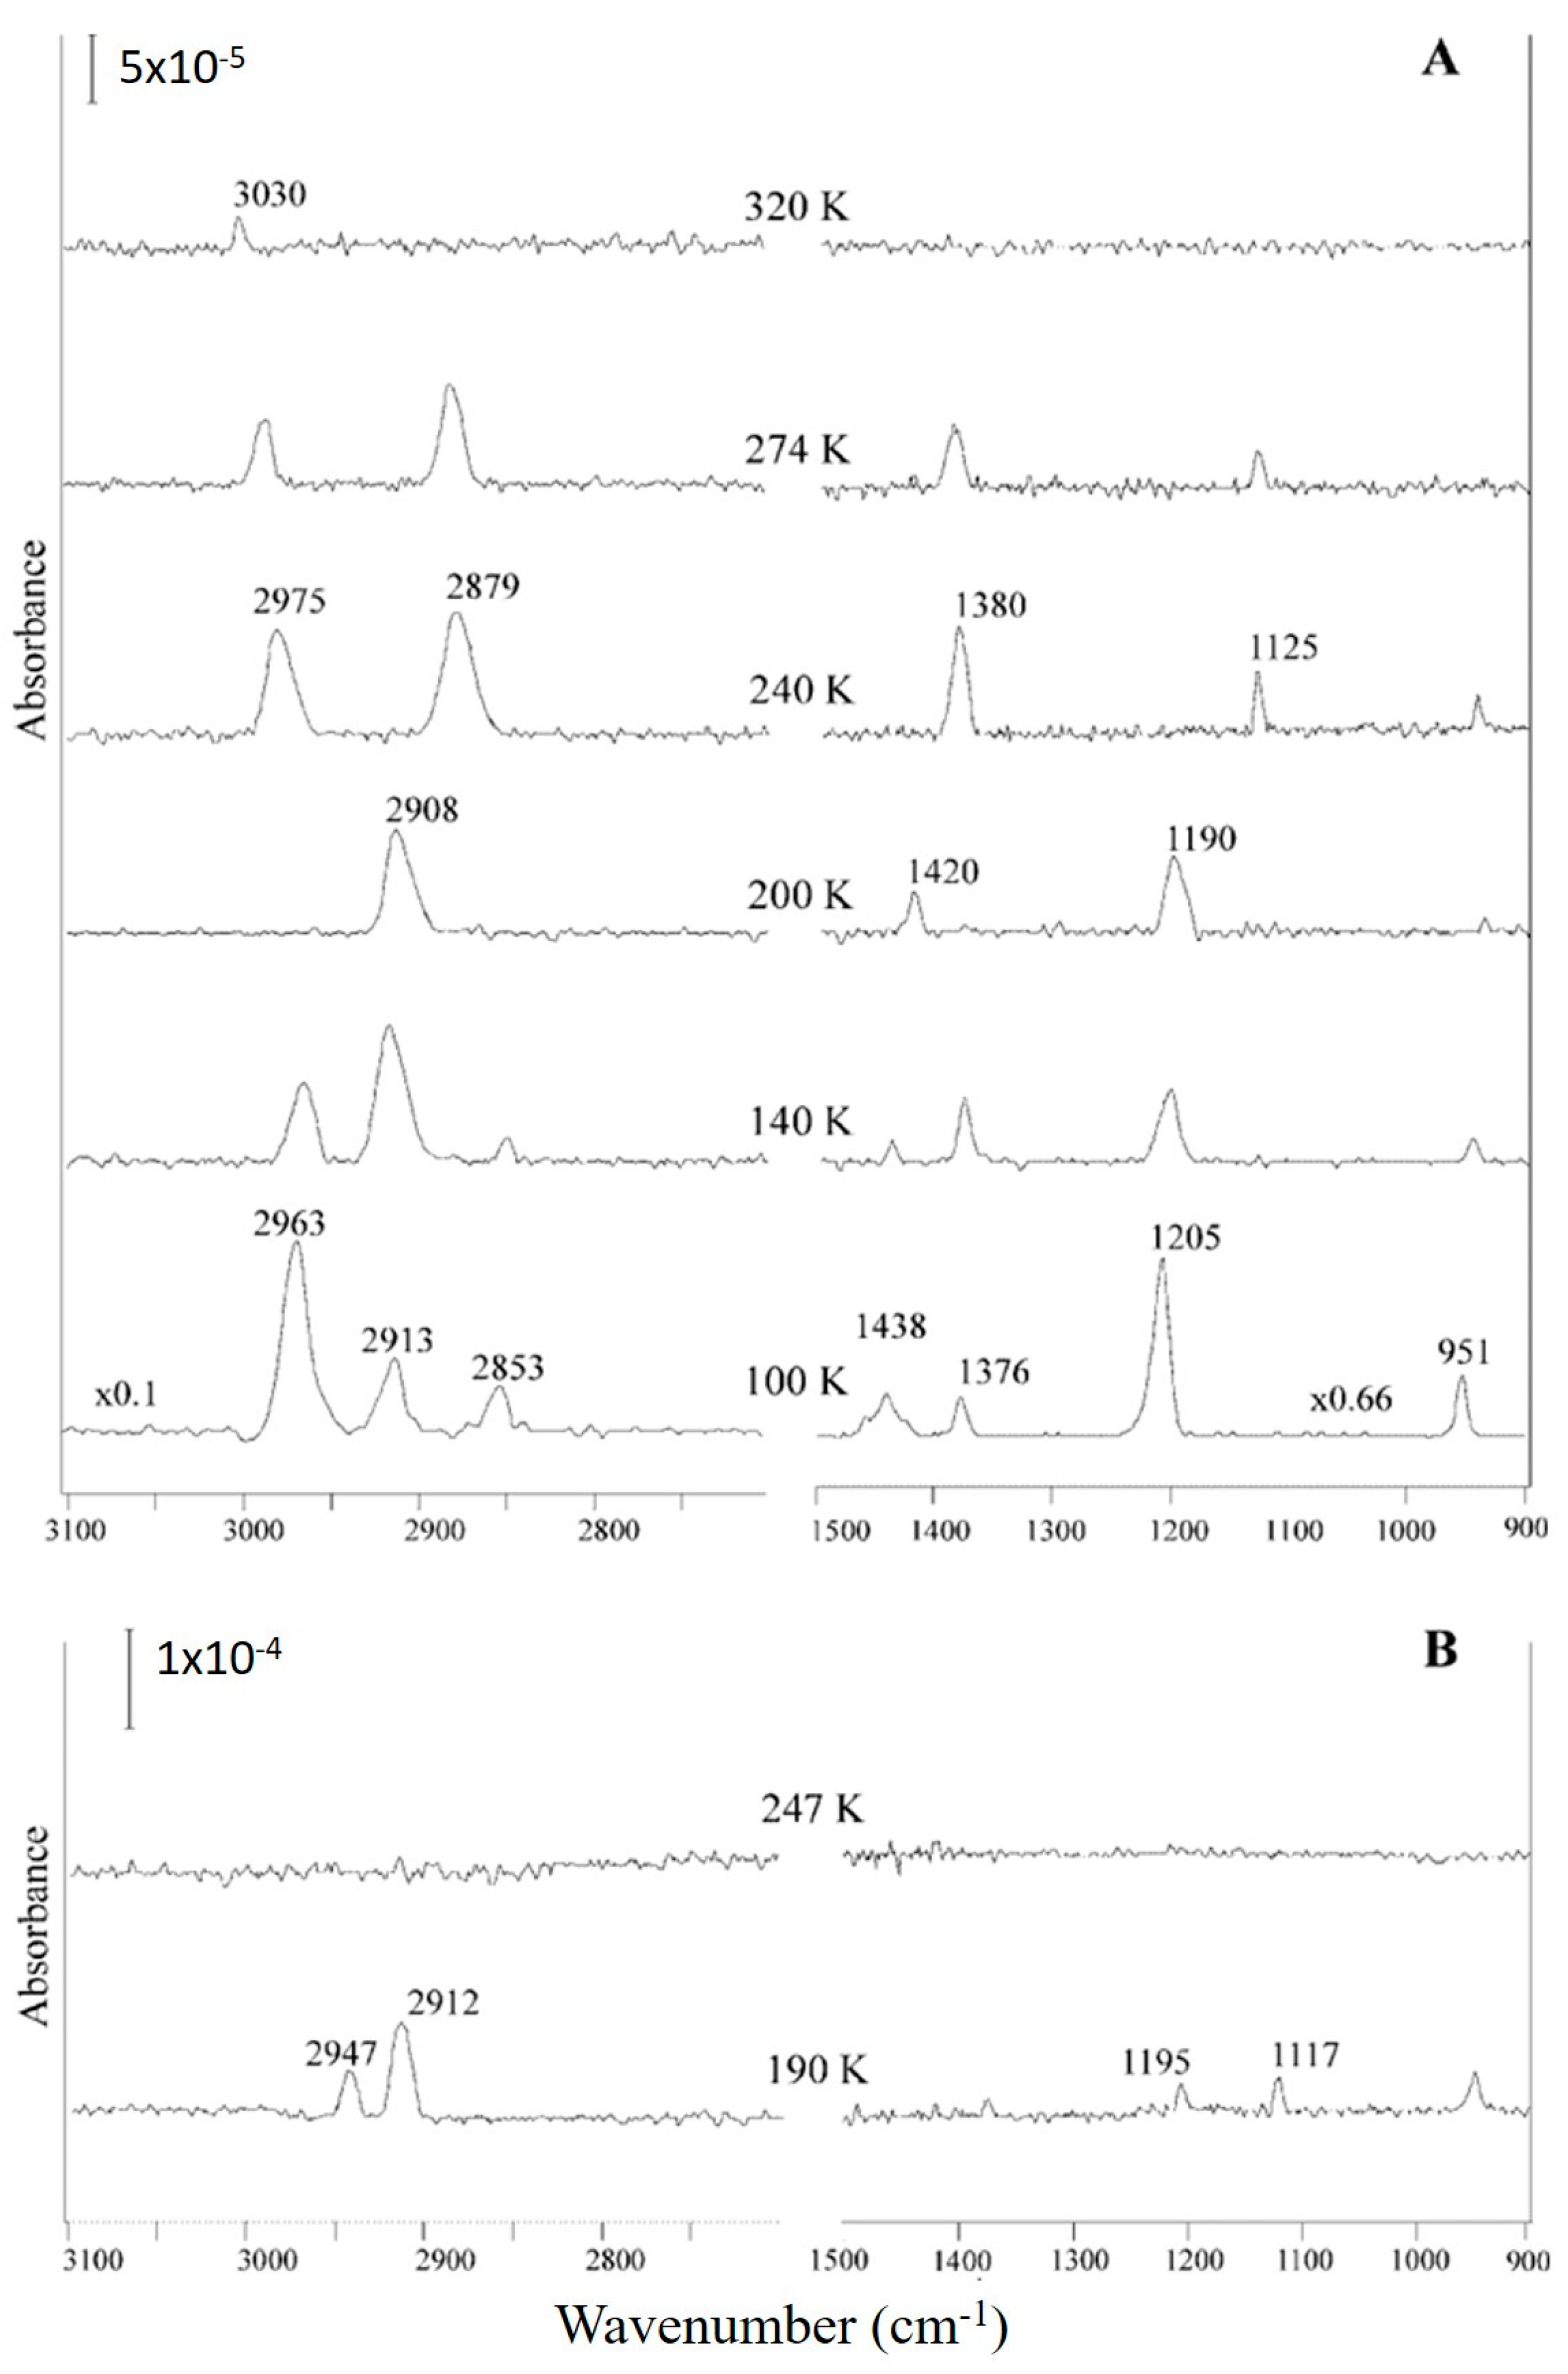
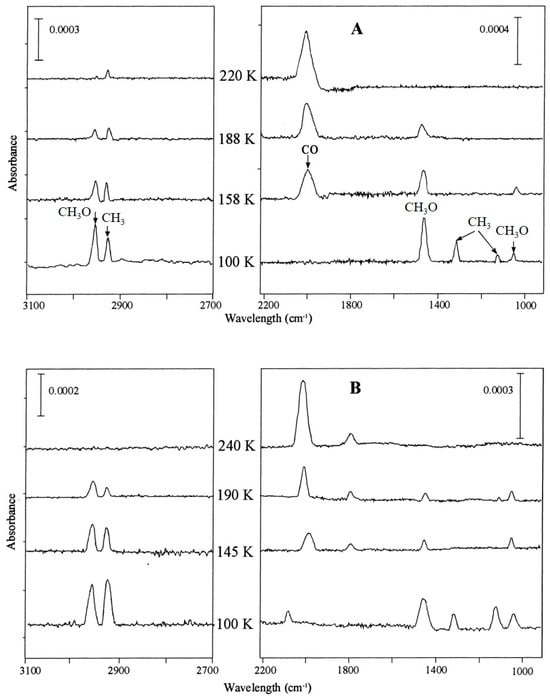
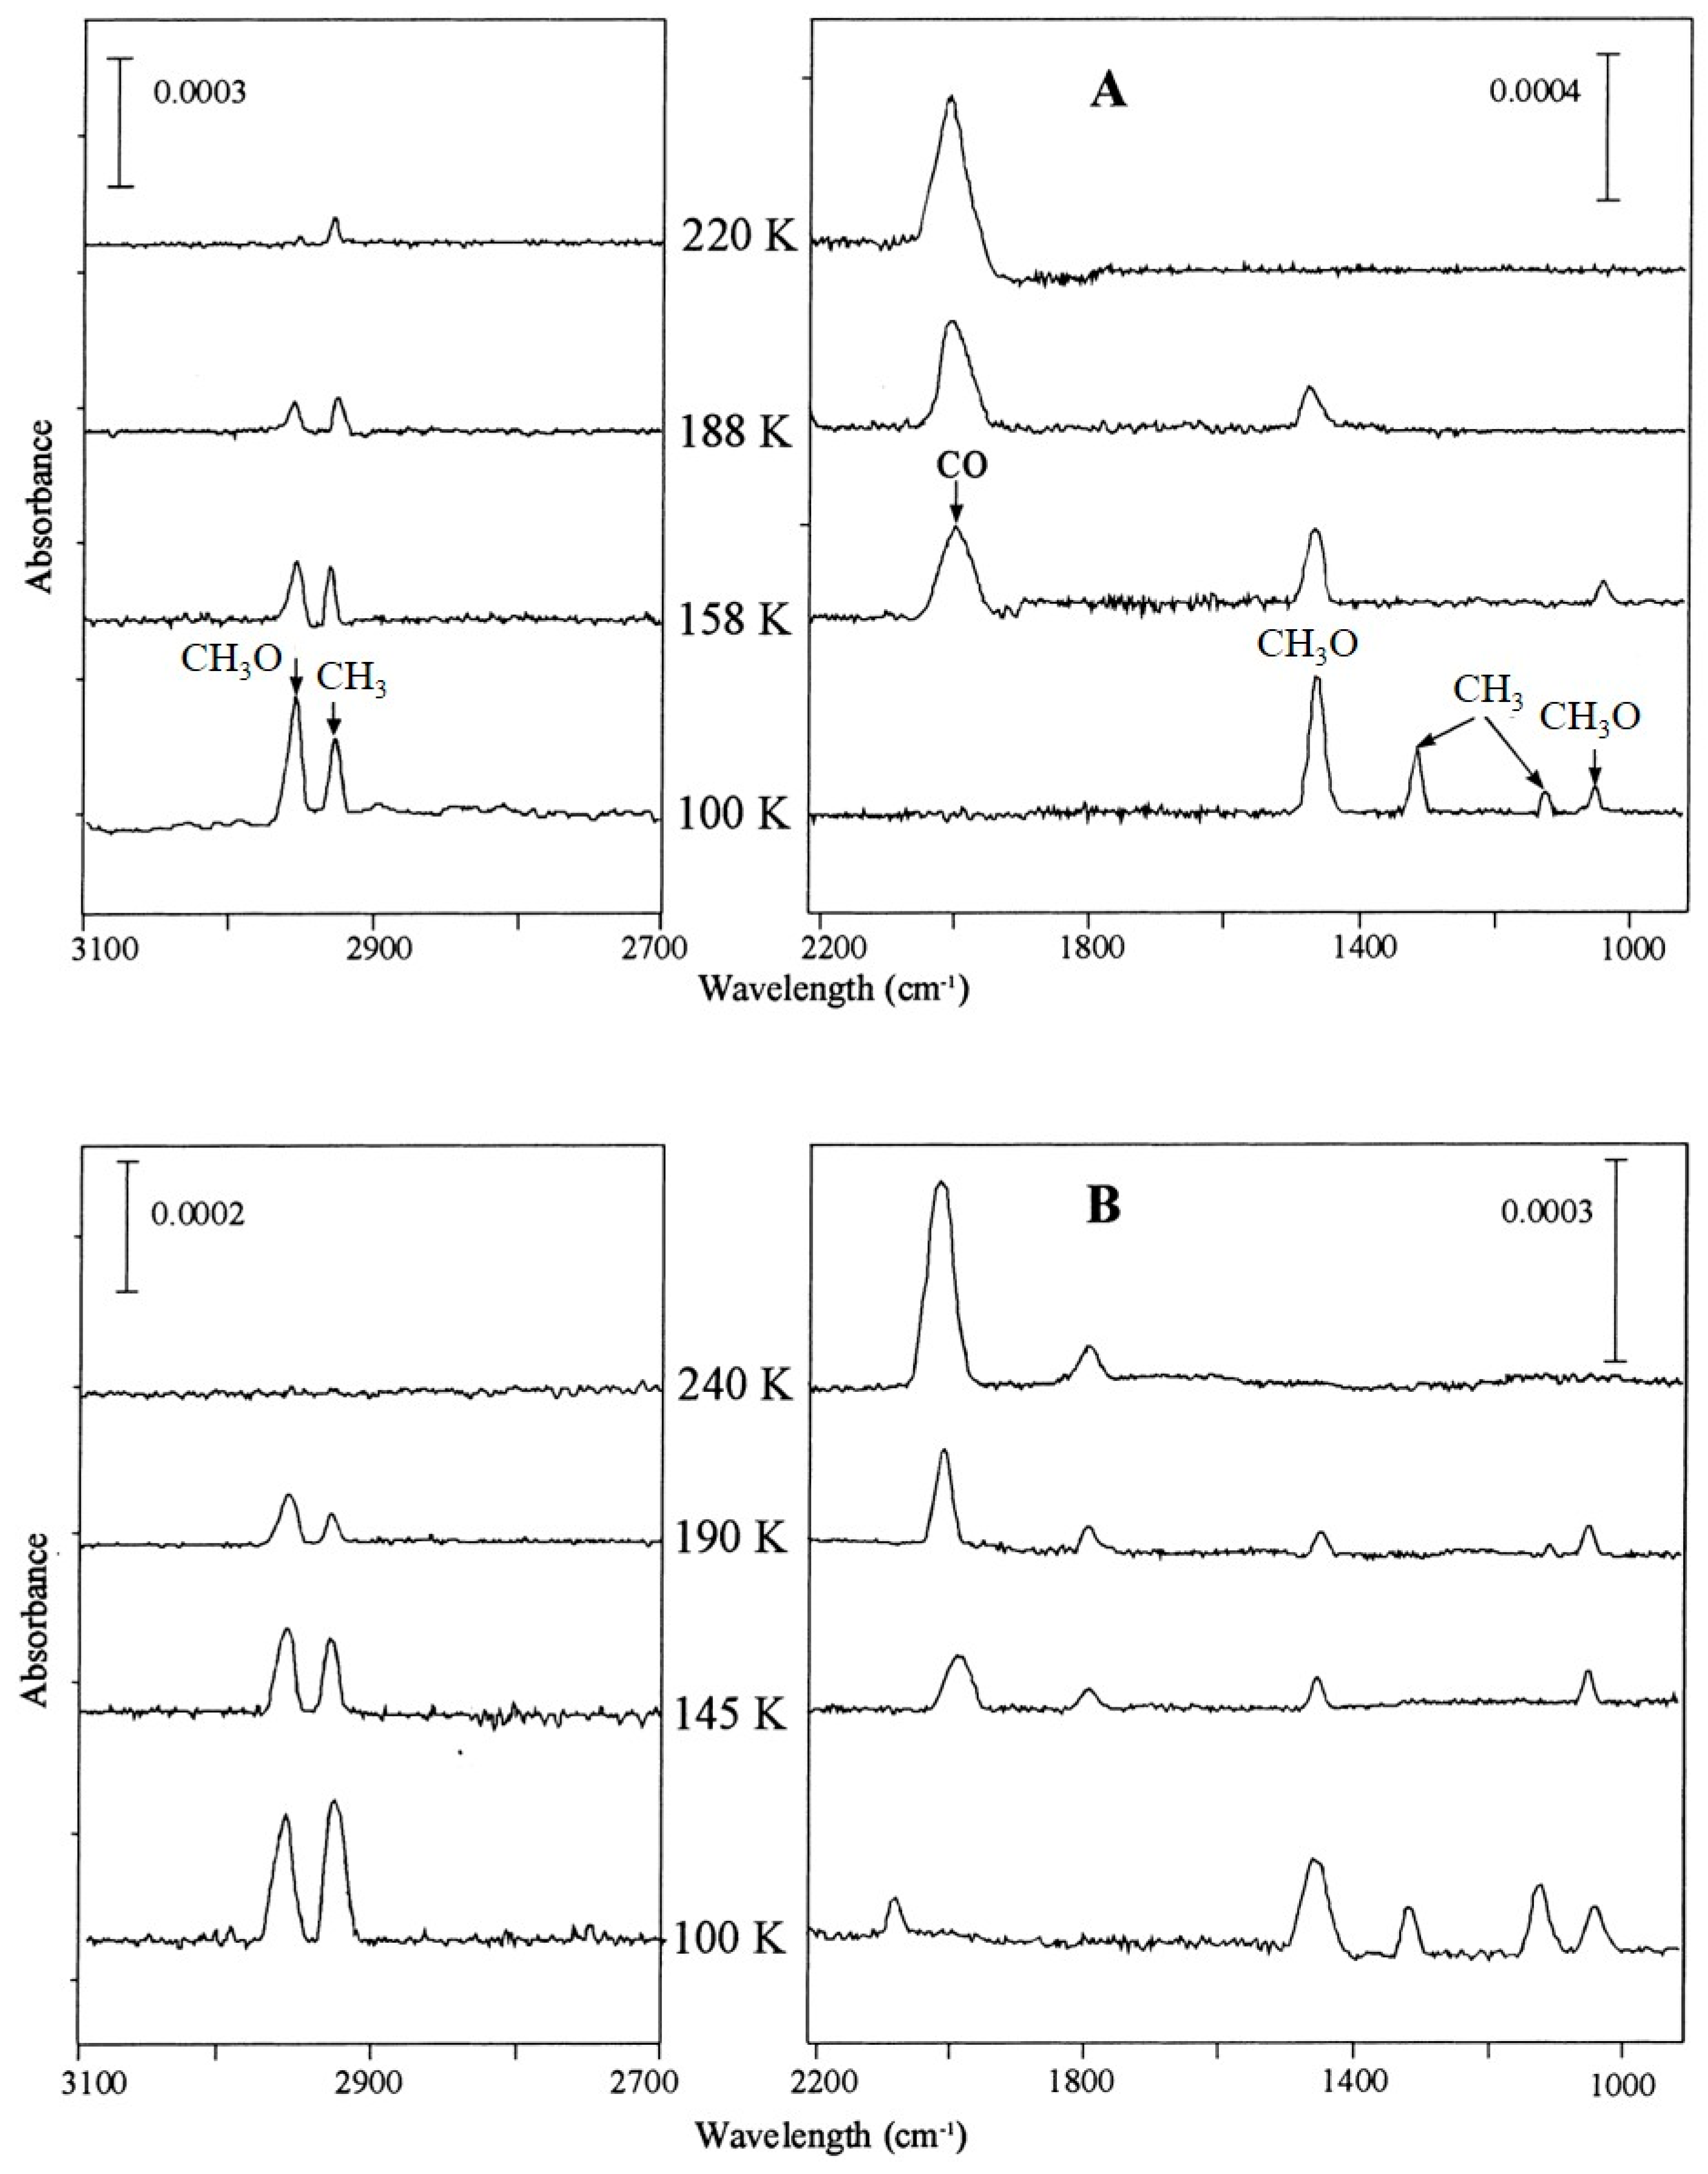
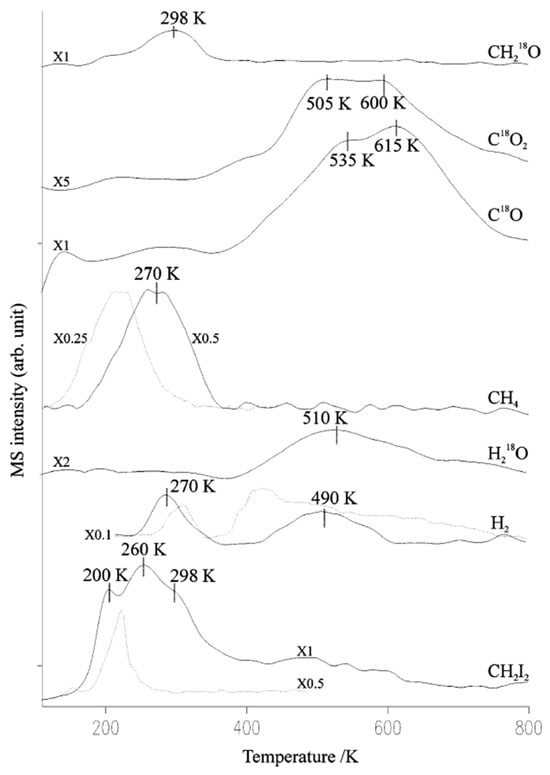
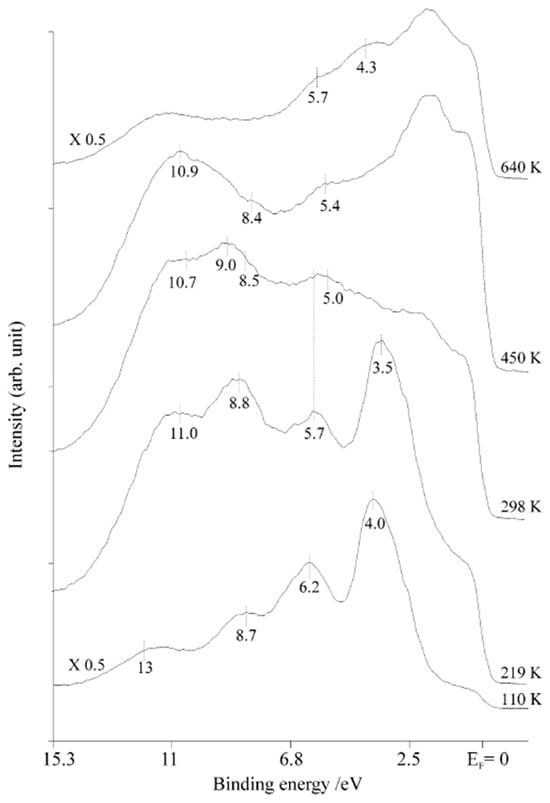
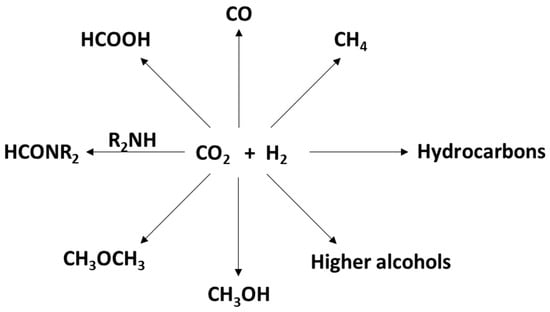
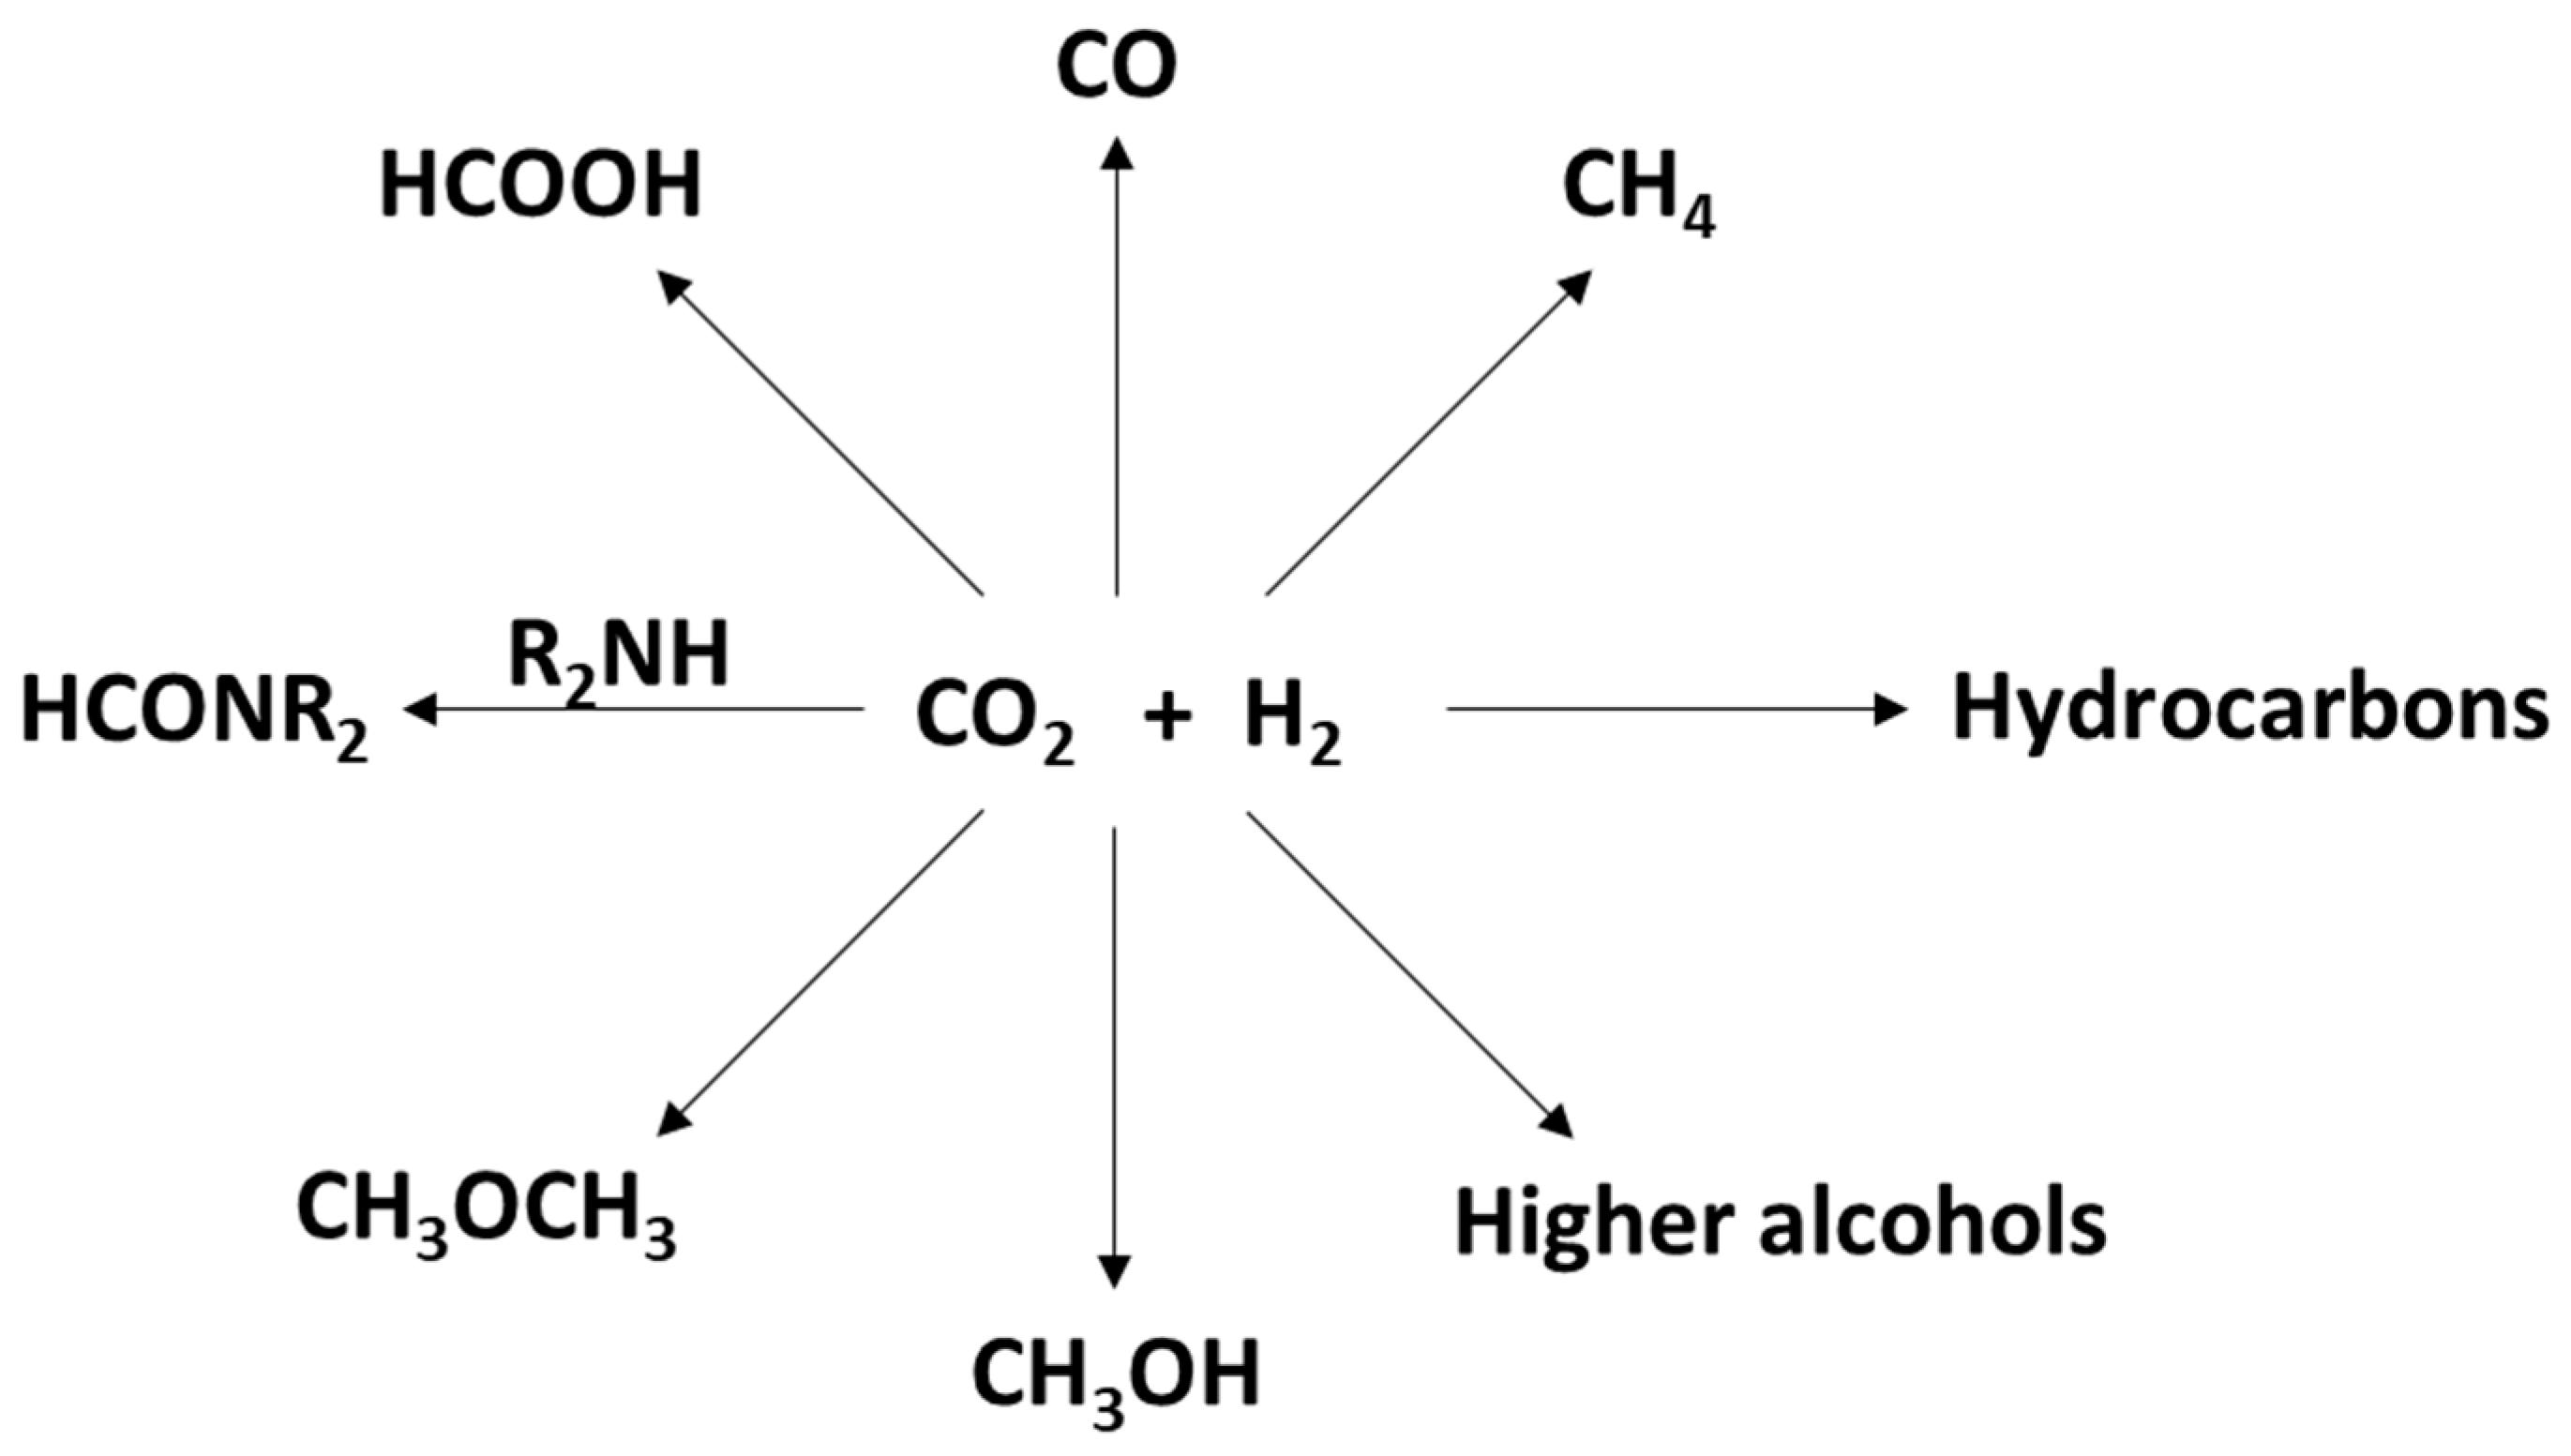

Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2026 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license.