


Abstract
Accurate and real-time state of charge (SOC) monitoring is critical for the safe, efficient, and stable long-term operation of vanadium redox flow batteries (VRFBs). Traditional monitoring methods are susceptible to errors arising from side reactions, cumulative drift, and electrolyte imbalance. This study develops a non-invasive optical sensor module for the negative electrolyte (anolyte), utilizing the favorable spectral properties of V(II)/V(III) ions at 850 nm for real-time SOC tracking. A fifth-order polynomial model was employed for calibration, successfully managing the non-linear optical response of highly concentrated electrolytes and achieving exceptional accuracy (adjusted R2 > 0.9999). The optical sensor reliably tracked capacity degradation over 50 galvanostatic cycles, yielding a degradation curve that showed a high correlation with the conventional coulomb counting method, thus confirming its feasibility for assessing battery’s state of health. Contrary to initial expectations, operating at higher current densities resulted in a lower capacity degradation rate (CDR). This phenomenon is primarily attributed to the time-dependent nature of parasitic side reactions. Higher current densities reduce the cycle duration, thereby minimizing the temporal exposure of active species to degradation mechanisms and mitigating cumulative ion imbalance. This mechanism was corroborated by physicochemical analysis via UV-Vis spectroscopy, which revealed a strong correlation between the severity of spectral deviation and the CDR ranking. This non-invasive optical technology offers a low-cost and effective solution for precise VRFB management and preventative maintenance.
1. Introduction
With the increasing global demand for renewable energy, the large-scale integration of intermittent energy sources such as solar and wind power poses significant challenges to grid stability [1]. Large-scale energy storage systems are considered a key technology to address this issue [1,2]. Among the numerous storage solutions, the all-vanadium redox flow battery (VRFB) has emerged as a highly promising candidate for medium-to-large stationary storage applications due to its long cycle life, high safety, independent scalability of power and capacity, and recyclable electrolyte [3,4,5]. In a VRFB, both the positive and negative electrolytes contain only vanadium species in an aqueous sulfuric acid solution, which effectively avoids cross-contamination resulting from crossover through the ion-exchange membrane, a major advantage over other flow battery types.
To ensure the safe, efficient, and long-term stable operation of VRFB systems, accurate, real-time monitoring of the state of charge (SOC) is crucial [6,7]. Traditional SOC monitoring methods each have their limitations. Moreover, mitigating parasitic side reactions [8], such as the hydrogen evolution reaction (HER) at the negative electrode and air oxidation of V(II) ions, is essential for maintaining capacity and longevity. Capacity maintenance strategies encompass both active surface modification techniques and critical pretreatment steps. Recent advanced strategies focus on actively suppressing HER by modifying electrode surfaces [9], for instance, through the electrochemical deposition of bismuth or other metals to shift the HER overpotential. Furthermore, high-performance VRFB systems typically utilize electrodes that undergo intensive pretreatment, such as thermo-chemical activation of carbon felt, involving immersion in concentrated sulfuric acid and subsequent oxidation at 400 °C. To counter V2+ oxidation by atmospheric oxygen, standard operational practice includes purging the negative half-cell electrolyte with high-purity nitrogen (N2) and covering the electrolyte surface with paraffin oil prior to operation. The open-circuit voltage method is accurate only when the system is balanced and cannot independently assess the state of the two half-cells [10]; if the system becomes imbalanced due to asymmetric ion crossover or side reactions (e.g., air oxidation of V(II), hydrogen evolution), the OCV method will not accurately reflect the SOC. Coulomb counting (current integration) is simple but susceptible to cumulative errors from current measurement and side reactions, leading to decreased accuracy, especially over long-term operation. Other direct methods also present significant trade-offs regarding cost, accuracy, and practicality. The conductivity method is highly sensitive to changes in temperature and electrolyte composition, requiring frequent calibration [10]. Potentiometric titration is precise but involves complex, time-consuming procedures, making it unsuitable for online, real-time monitoring [6]. More recently, advanced direct measurement techniques have been investigated [8]. Acoustic analysis methods estimate SOC by measuring ultrasonic velocity, showing high consistency and low error (e.g., ±2%) within the 10–90% SOC range. However, this approach is highly sensitive to temperature fluctuations, often necessitating high-precision temperature compensation. Measuring the acoustic attenuation coefficient offers lower accuracy (maximum error 4.8%) but is less temperature-sensitive. Amperometric methods, utilizing novel sensors (e.g., microelectrodes) to measure the limiting mass transfer current, can achieve very high accuracy (e.g., R2 = 0.9979). Nevertheless, they typically require expensive additional sensor components, increasing instrumentation costs. Electrolyte property-based estimation methods include measuring viscosity and density. While viscosity sensors are relatively inexpensive and easy to integrate, the measurements are greatly affected by temperature. Density-based methods perform better but rely on expensive measuring instruments and require calibration.
Therefore, developing a method that can independently, instantly, and accurately monitor the SOC of each half-cell is essential for the timely detection and remediation of electrolyte imbalance, thereby enabling the restoration of battery capacity [11]. In recent years, optical spectroscopic analysis has become a prominent research area for VRFB SOC monitoring due to its non-invasive nature, high sensitivity, rapid response, and potential for online, real-time (in-operando) implementation [10]. Its fundamental principle relies on the distinct colors and absorption spectra exhibited by vanadium ions in different valence states during charge–discharge cycles. In the negative electrolyte, green trivalent vanadium ions (V3+) are reduced to violet divalent vanadium ions (V2+) during charging; in the positive electrolyte, blue tetravalent vanadium ions (VO2+) are oxidized to yellow pentavalent vanadium ions (VO2+). By measuring the changes in absorbance or transmittance of the electrolyte at specific wavelengths, the relative concentrations of each valence state of vanadium can be quantitatively analyzed, allowing for the independent calculation of the SOC for both the positive and negative half-cells [7].
While the principles of spectroscopy are intuitive, applying them to commercial VRFB systems presents significant challenges. Commercial systems typically require high vanadium concentrations (≥1.5 M) to achieve competitive energy densities [6]. At these concentrations, the electrolyte behaves as a non-ideal solution dominated by strong intermolecular interactions, causing severe deviations from the linearity predicted by the Beer–Lambert law. This non-linearity complicates direct quantitative analysis and necessitates robust calibration models, Consequently, a high-order polynomial fitting model, such as the employed fifth-order polynomial model, is necessary to accurately capture this complex physical deviation, successfully managing the non-linear optical response of highly concentrated electrolytes and achieving exceptional accuracy (adjusted R2 > 0.9999) across different batches. Direct measurement of the concentrated stock solution can lead to signal saturation or complete opacity [1,10]. Consequently, many studies have resorted to diluting the electrolyte before measurement, a practice unsuitable for online, real-time monitoring [2]. To solve this problem, the most direct and effective method is to significantly shorten the optical path length [12] by using specially designed flow cells or quartz cuvettes with path lengths reduced to the millimeter or even micrometer scale (e.g., 0.5 mm, 0.2 mm, 0.1 mm, 50 µm). This allows the absorbance to be controlled within the effective measurement range of the instrument without diluting the sample.
Furthermore, the spectral behavior of the positive electrolyte (catholyte) is inherently complex [10]. The coexistence of V(IV) and V(V) ions leads to the formation of a strongly absorbing 1:1 mixed-valence complex, nominally V2O3 [12]. This formation results in “excess absorbance,” where the total optical density significantly exceeds the linear sum of the individual components, creating a non-linear, parabolic relationship between absorbance and SOC that introduces ambiguity in measurement [13]. Consequently, the relationship between catholyte absorbance and SOC becomes non-linear and parabolic, creating ambiguity in SOC determination [1]. This means that a single absorbance value can correspond to two different SOC values, creating ambiguity in accurate SOC determination [7]. To overcome this issue, researchers [11] have proposed several solutions: one is to use the entire spectral shape as a fingerprint and compare it against a pre-established database of standard spectra at various SOCs and concentrations, which is accurate but requires extensive preliminary calibration; another [13] is to deconvolve the complex measured spectrum into the contributions of its individual components (V(IV), V(V), and the complex); a more practical approach [14] is to accept the non-linear relationship and establish a calibration curve through mathematical fitting (e.g., with a second- or higher-order polynomial). Additionally, some studies [15,16] have explored the near-infrared region (e.g., 950 nm) for a more monotonic response or have suggested simultaneous measurements at two different wavelengths to resolve the ambiguity [17].
Furthermore, traditional laboratory spectrophotometers require a reference blank solution with the same solvent composition as the sample to calibrate for background absorption. In a dynamically operating VRFB system, the electrolyte composition may change over time, making it difficult to prepare an accurately matched reference solution. To address this, Shin et al. [18] introduced an innovative concept of difference absorbance. The core idea is to use the spectrum of the electrolyte at a known initial state (e.g., 0% SOC) as the reference spectrum, with subsequent absorbance measurements calculated as the change relative to this initial state. This method effectively eliminates the influence of the solvent and the system’s background absorption without requiring an additional reference fluid path, thereby greatly simplifying the design of an online monitoring system.
In contrast to the catholyte, the negative electrolyte (anolyte, V(II)/V(III) system) exhibits a simpler, near-linear spectral response in the near-infrared region. Building on this advantage, this study develops a non-invasive optical sensor module specifically targeting the anolyte. By utilizing a specific wavelength of 850 nm—selected to balance absorbance intensity with dynamic range—and integrating a fifth-order polynomial calibration to account for concentration effects, we propose a technically feasible solution for precise, real-time SOC tracking without the need for sample dilution. Because the electrolyte exhibits a non-linear trend at higher concentrations, this phenomenon cannot be adequately described by the simple Beer–Lambert law. This is due to the severe deviation from the linear relationship at high concentrations where intermolecular interactions become significant. In practice, employing a polynomial model for regression is preferred to accurately establish the relationship between the optical sensor and the SOC. Although V(II) ions are highly susceptible to air oxidation [10], this issue can be effectively managed in a sealed, online monitoring system [12,18].
The proposed non-invasive optical sensor module utilizes the favorable spectral properties of V(II)/V(III) ions to offer a robust solution. Crucially, this method allows for the independent determination of the anolyte state, providing insights into ion imbalance that are often masked in varying voltage or coulomb counting methods. The proposed optical sensor was validated over 50 galvanostatic cycles, yielding a degradation curve that highly correlates with conventional methods, thereby confirming its feasibility for assessing the battery’s state of health (SOH). Combined with technologies that simplify the system, such as ultra-short path-length flow cells and the difference absorbance method, it promises to provide an economical and reliable solution for the intelligent operational management of VRFBs.
2. Principle of VRFB and Its Challenges of Optical Monitoring
2.1. Principle of VRFB and Definition of SOC
VRFB is an electrochemical system that stores and releases energy through reversible redox reactions between different valence states of vanadium ions. During the charging process, tetravalent vanadium ions (V4+, VO2+) in the positive electrolyte (catholyte) are oxidized to pentavalent vanadium ions (V5+, VO2+), while trivalent vanadium ions (V3+) in the negative electrolyte (anolyte) are reduced to divalent vanadium ions (V2+). The reverse reactions occur during discharge. The electrochemical reactions are as follows:
SOC is a critical parameter for assessing the remaining capacity of a battery, defined as the ratio of the concentration of active species to the total concentration. For a VRFB, the SOC of the positive and negative half-cells can be defined as:
where represents the concentration of each vanadium species. Accurate and real-time SOC monitoring is crucial for the battery management system to manage issues such as self-discharge, capacity loss due to ion crossover, and electrode corrosion caused by high V5+ concentrations.
2.2. Challenges of Optical Monitoring
Traditional SOC detection methods based on voltage and current measurements are unable to effectively address the impacts of dynamic operation or cumulative errors. Therefore, this study employs ultraviolet–visible (UV-Vis) spectrophotometry, a method based on physicochemical properties, which is advantageous for its non-invasive nature, high sensitivity, and rapid response. The principle relies on the unique colors and optical absorption spectra of vanadium ions in different valence states. Geiser et al. [14] utilized an in-situ system for simultaneous measurement of open-circuit potential and absorption spectra. The characteristic UV-Vis absorption peaks for aqueous vanadium ions are distinct for each oxidation state. The violet V2+ ion exhibits two main absorption bands, one centered at approximately 570 nm and a broader band in the near-infrared region of 850–950 nm. The green V3+ ion is characterized by absorption maxima at around 400 nm and 600 nm. In the +4 oxidation state, vanadium exists as the blue vanadyl cation, VO2+, which displays a prominent absorption peak in the 760–780 nm range. Finally, the yellow pervanadyl ion, VO2+, representing the +5 oxidation state, shows strong absorption at the blue end of the visible spectrum, typically between 400–430 nm, with another significant absorption band in the ultraviolet region below 350 nm.
However, direct application of spectrophotometry to commercial-grade, high-concentration VRFB electrolytes faces two major challenges. The first is the limitation of the Beer–Lambert law; the high absorbance of concentrated electrolytes leads to signal saturation, preventing direct measurement. Research by Loktionov et al. [13] indicated that a 2.0 M vanadium electrolyte required a 20-fold dilution for full-spectrum analysis. The second challenge is the complex spectral behavior of the catholyte. At a wavelength of 754 nm, its absorbance exhibits a non-linear, parabolic relationship with SOC, and its light absorption is excessively strong over a wide SOC range (5–80%), making it unsuitable for direct measurement with a photodiode. Additionally, electrolyte composition parameters, specifically the sulfuric acid concentration (and associated H+ concentration or pH), fundamentally affect both the electrochemical performance and the resulting UV-Vis spectral properties. Changes in sulfuric acid concentration influence the electrolyte conductivity [10], the thermodynamic calculation of SOC via the Nernst potential [14], and the vanadium ion speciation (particularly affecting the V(V) species and complex formation in the positive electrolyte) [11]. Therefore, for optical monitoring techniques, the dependence of absorbance on the supporting electrolyte concentration must be quantified and modeled, especially when aiming for accurate SOC readings across different electrolyte batches.
In contrast, the spectral properties of the anolyte (V(II)/V(III) system) in the near-infrared (NIR) region are more favorable [11]. While absorbance is strong at the primary characteristic peaks of V3+ (400 nm and 600 nm) making them susceptible to the Beer-Lambert law saturation limits when measured without dilution, the change in absorbance is more manageable in the NIR region. Quantitative analysis of the anolyte revealed distinct performances between 750 nm and 850 nm. At 750 nm, the absorption intensity variation was observed to be moderate, ranging from 0.2 to 0.8 (increasing with SOC). Conversely, at 850 nm, a significantly wider dynamic range was observed, with absorption intensity variation ranging from 0 to 1.3 (also increasing with SOC). Rudolph et al. [16] previously attempted to use a 950 nm infrared sensor for anolyte monitoring, while Geiser et al. [14] suggested using 440 nm. After a comprehensive evaluation, the 850 nm wavelength was selected for this study because the wider dynamic range (0 to 1.3) relative to other viable NIR options provides the optimal balance: moderate absolute absorbance levels coupled with a potential for higher resolution and signal-to-noise ratio over the full SOC range, making it suitable for developing a dilution-free, real-time monitoring sensor.
3. Materials and Experimental Methods
3.1. Materials and Experimental Apparatus
Three batches of all-vanadium electrolytes were used in this study. All samples had a total vanadium ion concentration ([V]) of 1.6 M but differed in their sulfate ([SO42−]) concentrations. The detailed specifications are provided in Table 1.
Table 1.
Batch numbers and concentrations of the all-vanadium electrolyte samples.
The experimental system consisted of a single-cell VRFB stack with an active electrode area of 25 cm2 (5 × 5 cm) and an electrolyte volume of 85 mL for each half-cell. The electrolyte was circulated by a GT-100 peristaltic pump operating at 40 rpm, corresponding to a stack outlet flow rate of 70 mL/min. Crucially, this study deliberately excluded standard electrolyte preservation and electrode activation protocols, such as thermo-chemical electrode pretreatment (e.g., concentrated sulfuric acid immersion and 400 °C oxidation) and negative electrolyte purging with high-purity N2 followed by paraffin oil coverage. This exclusion was mandated to accelerate the observation of capacity degradation rate (CDR) resulting from parasitic side reactions within the scope of 50 charge–discharge cycles, thereby obviating the necessity of testing over hundreds of cycles.
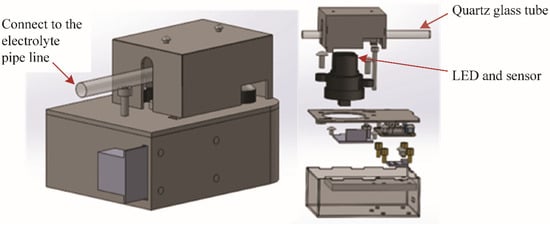
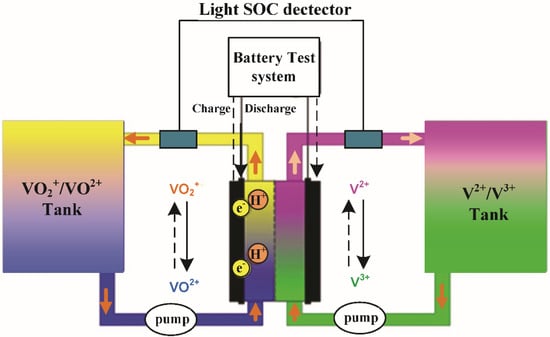
The core measurement device was an externally mounted optical SOC sensor module. This module was designed to clamp onto a quartz glass tube with a 6 mm inner diameter and integrates an infrared light-emitting diode (LED) source and a photodetector. Figure 1 shows the module and its exploded-view diagram. The initially selected wavelengths for the light source were 850 nm, 880 nm, and 890 nm. The sensor modules were installed on the outlet tubing of both the positive and negative electrodes of the stack to non-invasively monitor the optical properties of the electrolyte in real-time. The overall experimental setup is depicted in Figure 2.
Figure 1.
Schematic and exploded views of the optical SOC sensor module.
Figure 2.
Overall experimental setup showing the installation of the optical sensor modules.
Several instruments were used for characterization. A high-performance battery testing system (Neware 5V6A) was employed for galvanostatic charge–discharge cycling. A conductivity meter (Eutech CON 2700) was used to measure the electrolyte’s conductivity before and after cycling. Furthermore, a UV-Vis Spectrometer (PG Instruments T60) recorded the absorbance spectra of the electrolyte over a wavelength range of 300 to 1000 nm. For this measurement, samples were first diluted 20-fold with deionized water, which was also used as a blank to subtract the background.
3.2. Experimental Protocol and Data Analysis
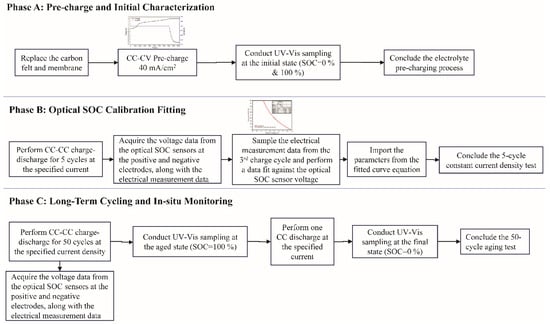
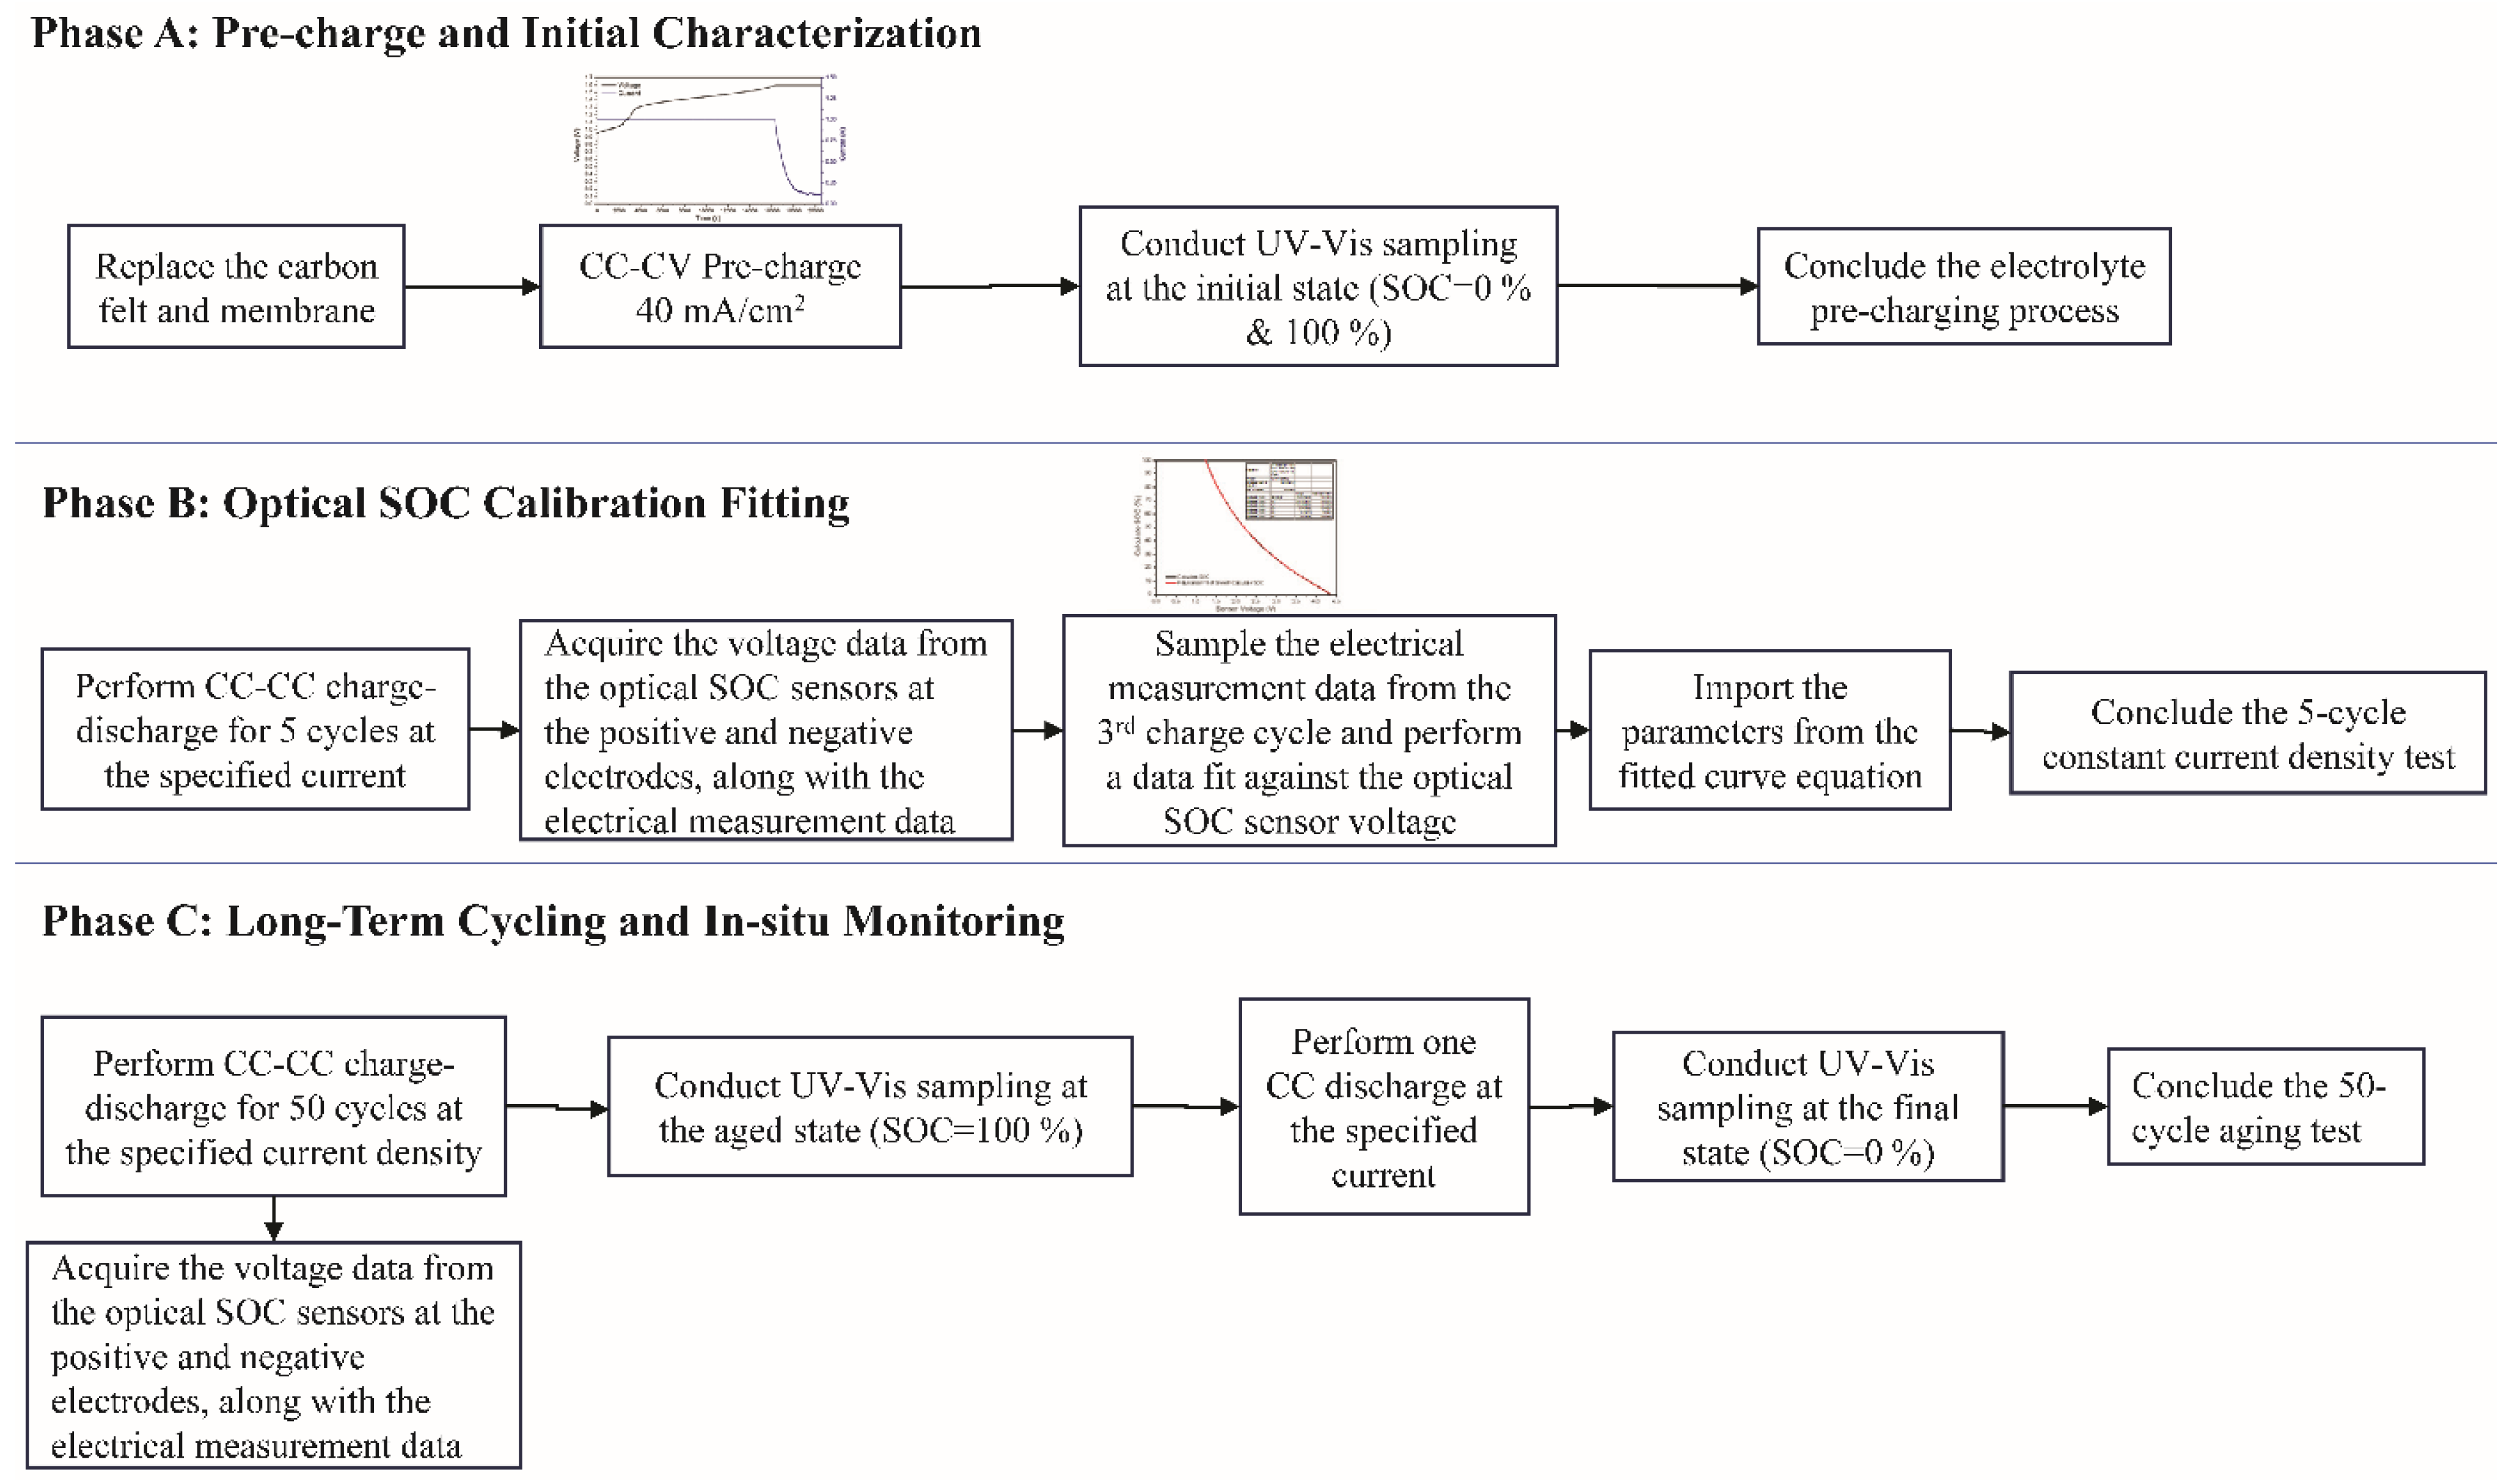
To ensure consistency across experiments, the proton exchange membrane and carbon felts were replaced before testing each new electrolyte. The experimental procedure, illustrated in the flowchart in Figure 3, was divided into three main phases.
Figure 3.
The overall experimental workflow for this study, illustrating the three phases of electrolyte testing and the respective characterization techniques employed.
- Phase A: Pre-charge and Initial Characterization
- Each half-cell was filled with 85 mL of electrolyte. An initial pre-charge was performed in constant current (CC) mode at 40 mA/cm2 up to 1.6 V, followed by a constant voltage (CV) step until the current dropped to a cutoff of 4 mA/cm2.
- After pre-charging (defined as SOC = 100%), small aliquots of the positive and negative electrolytes were sampled. Their UV-Vis spectra were analyzed after dilution. This constituted the Initial state data at SOC = 100%.
- The cell was then discharged at 40 mA/cm2 to a cutoff voltage of 0.7 V (defined as SOC = 0%). Samples were again collected and analyzed for UV-Vis spectra to obtain the Initial state data at SOC = 0%.
- Phase B: Optical SOC Calibration Fitting
- Following phase A, the cell was subjected to 5 galvanostatic charge–discharge cycles at 40 mA/cm2.
- During the third charging cycle, two datasets were recorded simultaneously: the raw voltage signal from the optical SOC sensor and the accumulated charge/discharge data from the battery cycler, which was used to calculate the SOC via the coulomb counting method.
- These two datasets were used to generate a fifth-order polynomial fitting curve, establishing the calibration equation between sensor voltage and SOC for that specific electrolyte batch. This equation was subsequently programmed into the sensor module to complete its calibration.
- Phase C: Long-Term Cycling and In-situ Monitoring
- Using the calibrated optical SOC modules, a 50-cycle lifetime test was conducted under four different constant current densities: 40, 60, 80, and100 mA/cm2.
- Throughout the 50 cycles, the SOC of the positive and negative electrolytes, as measured by the optical modules, was recorded in real-time, along with the electrochemical data from the battery cycler.
- Upon completion of 50 cycles (the final or aged state), the sampling procedure from Phase A was repeated at SOC = 0% and SOC = 100% to measure the final UV-Vis spectra for comparison with the initial state.
In addition, there are two important protocols to evaluate the reproducibility and stability of the proposed sensor system as follows.
3.2.1. Reproducibility Test
Due to the highly sensitive relationship between the measurement errors of the optical sensor and the coulomb counting method, and given that the resolution across different batches ranged between 0.14% and 0.44%, the error in repeated testing was primarily attributed to the experimental values obtained from the coulomb counting method. To establish measurement reliability, electrolyte batch #2306001 was used for 3 replicate experiments over 5 charge–discharge cycles, covering 15 conditions (3 current densities and 5 electrolyte flow rates).
3.2.2. Long-Term Stability Test
For evaluating the baseline stability, a long-term stability test was performed. The SOC value of electrolyte batch #2306001 was measured in a static (non-flowing) state. The drift rate was then calculated based on the 24 h of baseline drift data recorded by the sensor.
The electrochemical performance of VRFB was analyzed by the data from the battery cycler. They were used to calculate the coulombic efficiency (CE), voltage efficiency (VE), and energy efficiency (EE) for each cycle. Electrolyte utilization efficiency (UE) is a metric used to quantify the total amount of active species in the electrolyte that participates in electrochemical reactions. UE is calculated by the following formula:
The capacity degradation rate (CDR) is an indicator for evaluating the extent of decline in a battery’s ability to store energy after multiple charge–discharge cycles. This phenomenon, also termed capacity fading, is defined as the gradual reduction in the battery’s discharge capacity with an increasing number of cycles. A higher degradation rate value indicates a smaller remaining capacity after cycling. In this study, this rate is calculated by comparing the charge–discharge capacity of each cycle against that of the first cycle. The formula is as follows:
The polynomial model established in phase B was used to convert the real-time optical sensor voltage into SOC values. The SOC peak values from each cycle were plotted to track capacity fade and were compared with the results from the coulomb counting method.
Physicochemical characterization of the electrolyte was performed using UV-Vis spectroscopy. According to research by Geiser et al. [14], UV-Vis spectroscopic analysis of the positive and negative electrolytes can reveal characteristic peaks corresponding to the valence states of vanadium ions. In the positive electrolyte, peaks are generated for V5+ at <350 nm and for V4+ at 760 nm. The peak at <350 nm is less distinct as it is in the near-ultraviolet range and is susceptible to background interference. For the negative electrolyte, UV-Vis analysis at different battery SOC reveals four characteristic peaks at 400, 570, 600, and 850 nm. Among these, the 850 nm peak is difficult to distinguish under conditions of low variation and is prone to disappearing due to oxidation. Therefore, this study focuses on analyzing characteristic peaks at specific wavelengths: primarily the 760 nm peak representing V4+ in the positive electrolyte, the 400 nm and 600 nm peaks representing V3+ in the negative electrolyte, and the 570 nm peak representing V2+. By comparing the intensity changes in the 760 nm (V4+) and 400 nm (V3+) peaks and the wavelength shift (redshift) between 570 nm (V2+) and 600 nm (V3+) across various states before and after cycling, the degree of vanadium ion conversion and the extent of electrolyte imbalance before and after aging were quantitatively assessed.
4. Results and Discussion
4.1. Optical SOC Calibration Fitting and Resolution Analysis
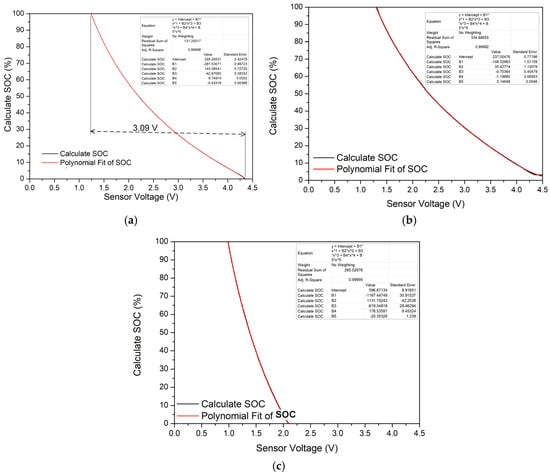
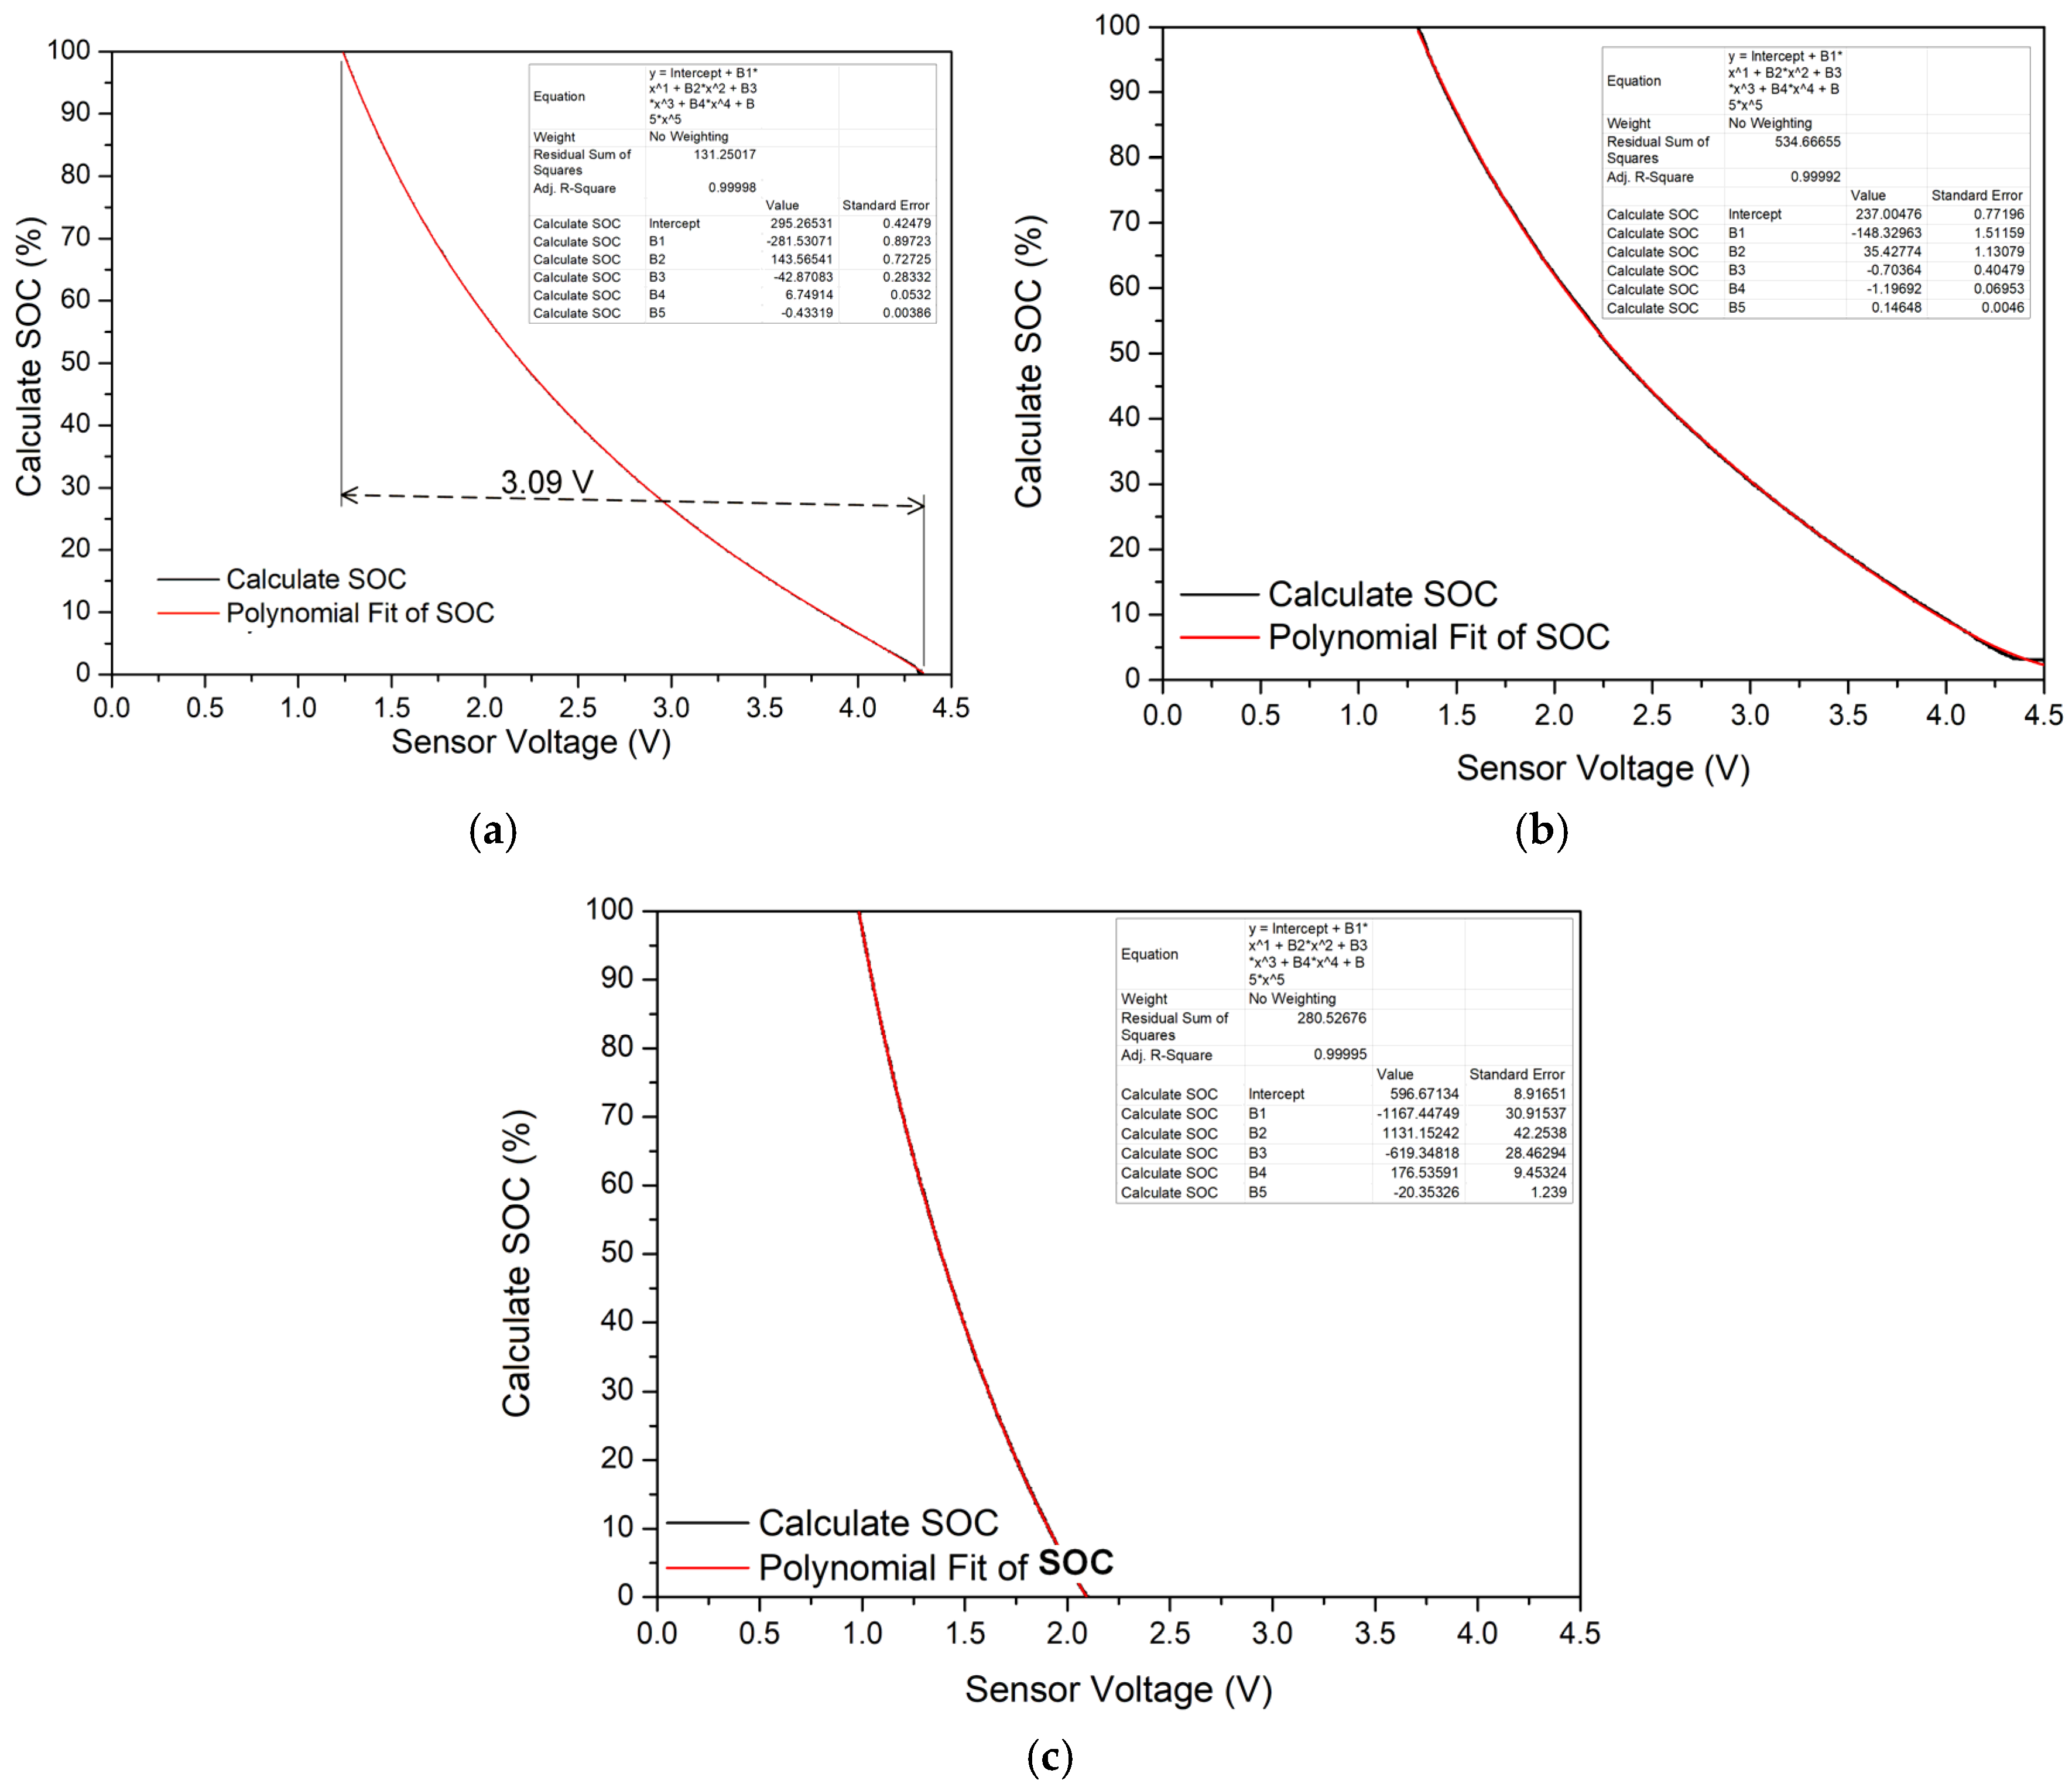
Prior to long-term cycling tests, it is essential to perform an optical SOC calibration for each electrolyte batch. In this study, following a pre-charge, the cell was subjected to 5 galvanostatic cycles at 40 mA/cm2, and data from the third charging cycle—specifically, the power data from the battery cycler (converted to SOC) and the raw voltage signal from the optical sensor- were used for fitting. A fifth-order polynomial model was employed to establish the calibration equation between the sensor voltage and SOC. In preliminary experiments, the anolyte data from 5 charge–discharge cycles using electrolyte batch #2306003 showed that polynomials of the fifth order or higher could achieve and maintain an accuracy of R2 = 0.99997 (see Figure S1). To prevent the use of higher-order polynomials from causing overfitting and to avoid increasing unnecessary computational load, the fifth-order model was thus used for fitting in this paper. The fitting results, illustrated in Figure 4, demonstrated excellent adjusted R2 values (>0.9999) for all batches, confirming that this model accurately describes the non-linear relationship between the two parameters.
Figure 4.
Optical SOC calibration fitting curves for the three electrolyte batches: (a) #2306001, (b) #2306002, and (c) #2306003.
The resolution of the sensor module is directly influenced by the optical property changes in the electrolyte. A larger change in the electrolyte’s optical properties during charging results in a greater voltage span from the sensor, which corresponds to a higher resolution for the optical SOC module. The analysis results, summarized in Table 2, highlight significant batch-to-batch variability in the optical response, which correlates strongly with the differences in sulfuric acid concentration. For instance, batch #2306002 (2.65 M H2SO4) exhibited the largest sensor voltage span of 3.51 V, yielding to the highest SOC resolution (0.14%). Conversely, batch #2306003 (2.9 M H2SO4) showed the smallest voltage span (1.11 V), resulting in the lowest resolution (0.44%). This pronounced variability in optical sensitivity underscores the fundamental finding that the electrolyte formulation, specifically the concentration of sulfuric acid, drastically modifies the spectral absorption characteristics of the V(II)/V(III) system at 850 nm. Consequently, batch-specific calibration of the fifth-order polynomial model is mandatory to ensure the module provides the specified high precision across different electrolyte supplies. The performance of batch #2306001 was intermediate, with a voltage span of 3.09 V and a resolution of 0.15%. These data indicate that the electrolyte formulation, particularly the sulfuric acid concentration, significantly affects the sensitivity of the optical response, thereby determining the precision of the optical monitoring. To rigorously assess measurement fidelity, a reproducibility test was conducted using batch #2306001 across 15 conditions (3 current densities and 5 electrolyte flow rates) for 3 replicate experiments over 5 cycles. The experimental data and estimation process are detailed in Table S1. The resultant standard deviation was 1.38%. It is estimated that when measuring under the same conditions using the optical sensor alone, the standard deviation for repeated experiments only marginally increases, falling between 1.38% and 1.44%, thereby faithfully reflecting the actual experimental error. Furthermore, a long-term stability test revealed that when measuring the SOC of batch #2306001 in a static state, the drift rate, calculated from 24 h of baseline drift data, was 0.509%.
Table 2.
Summary of optical SOC calibration fitting results for the three electrolyte batches.
Necessity of Batch-Specific Calibration and Limits of Generalization
The core findings from the calibration analysis demonstrate a strong inverse correlation between sulfuric acid concentration and the sensor’s optical response sensitivity (voltage span). The observed differences in sensor voltage spans (e.g., 3.51 V for Batch #2306002 vs. 1.11 V for Batch #2306003) exceed typical measurement tolerances, indicating that a calibration model derived from one electrolyte batch cannot be reliably generalized to monitor another batch with differing sulfuric acid concentrations. This absence of generalization capability mandates that the high-order polynomial model must be calibrated specifically for each distinct electrolyte batch. This batch-specific calibration step is essential for translating the variable optical signal into accurate SOC readings with high resolution (Adj. R2 > 0.9999). This requirement highlights the critical role of pre-use optical characterization in the deployment of this technology for VRFBs utilizing varying electrolyte compositions.
4.2. Electrolyte Utilization Under Different Current Densities
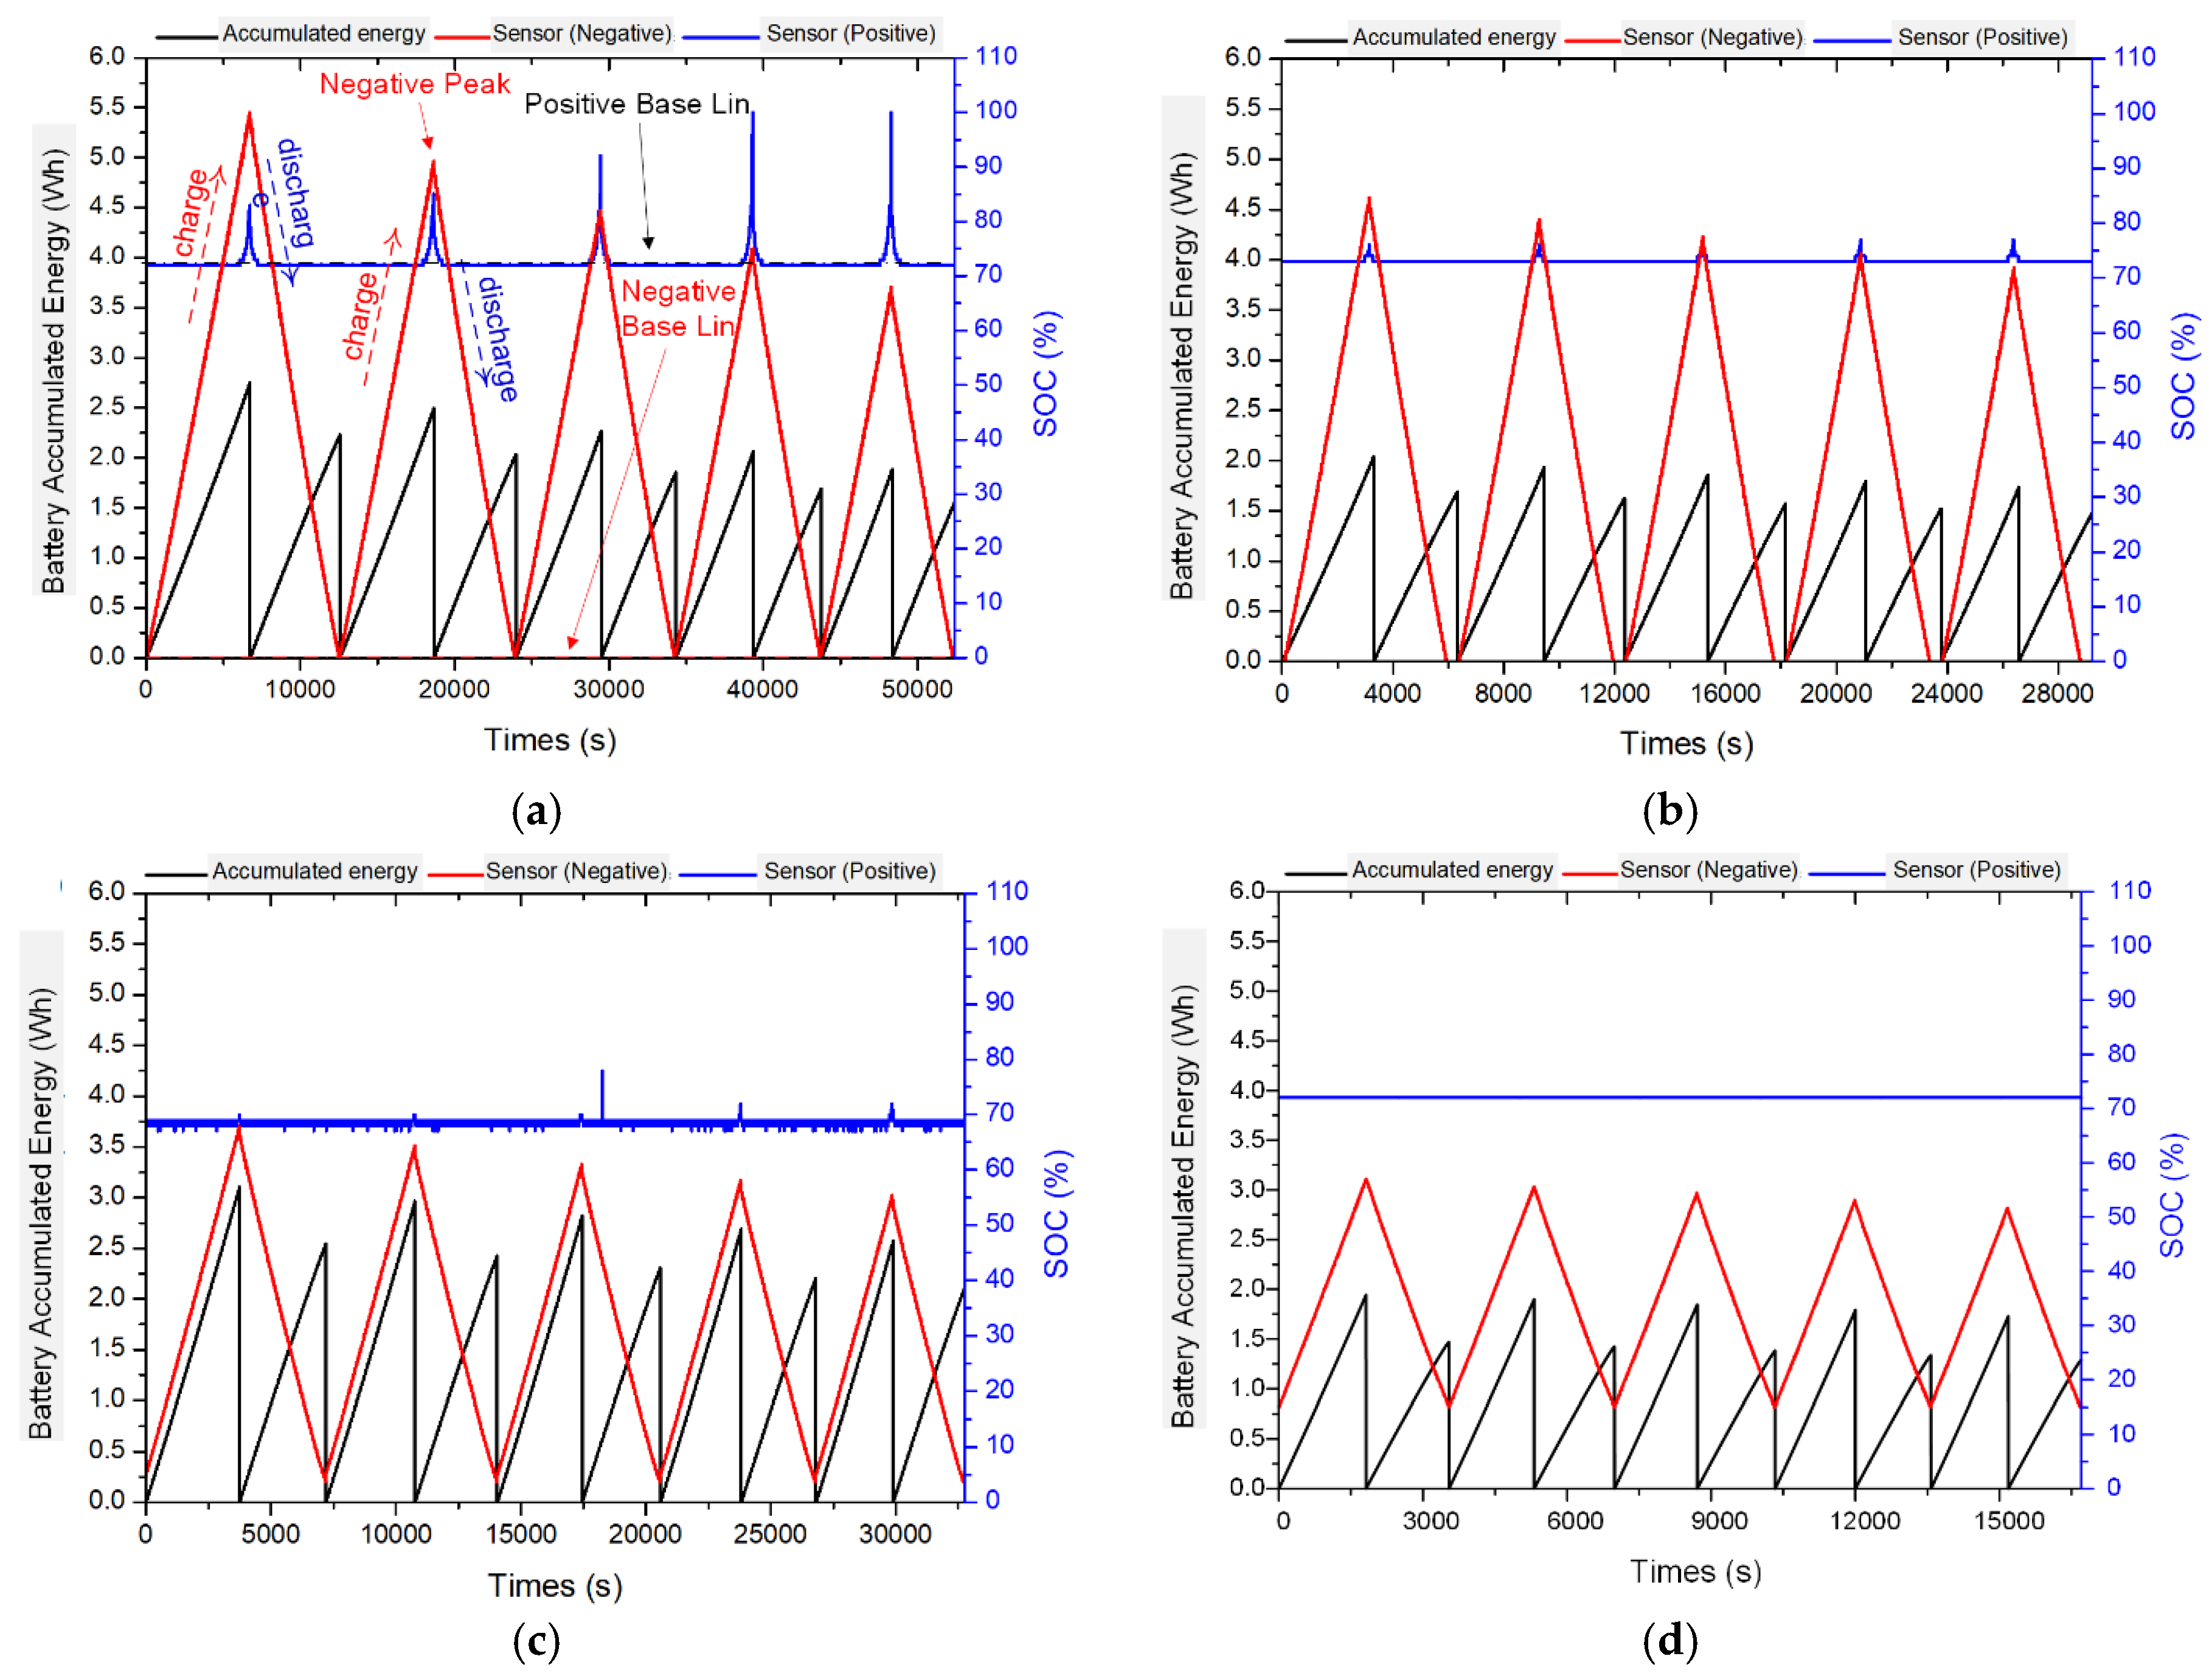
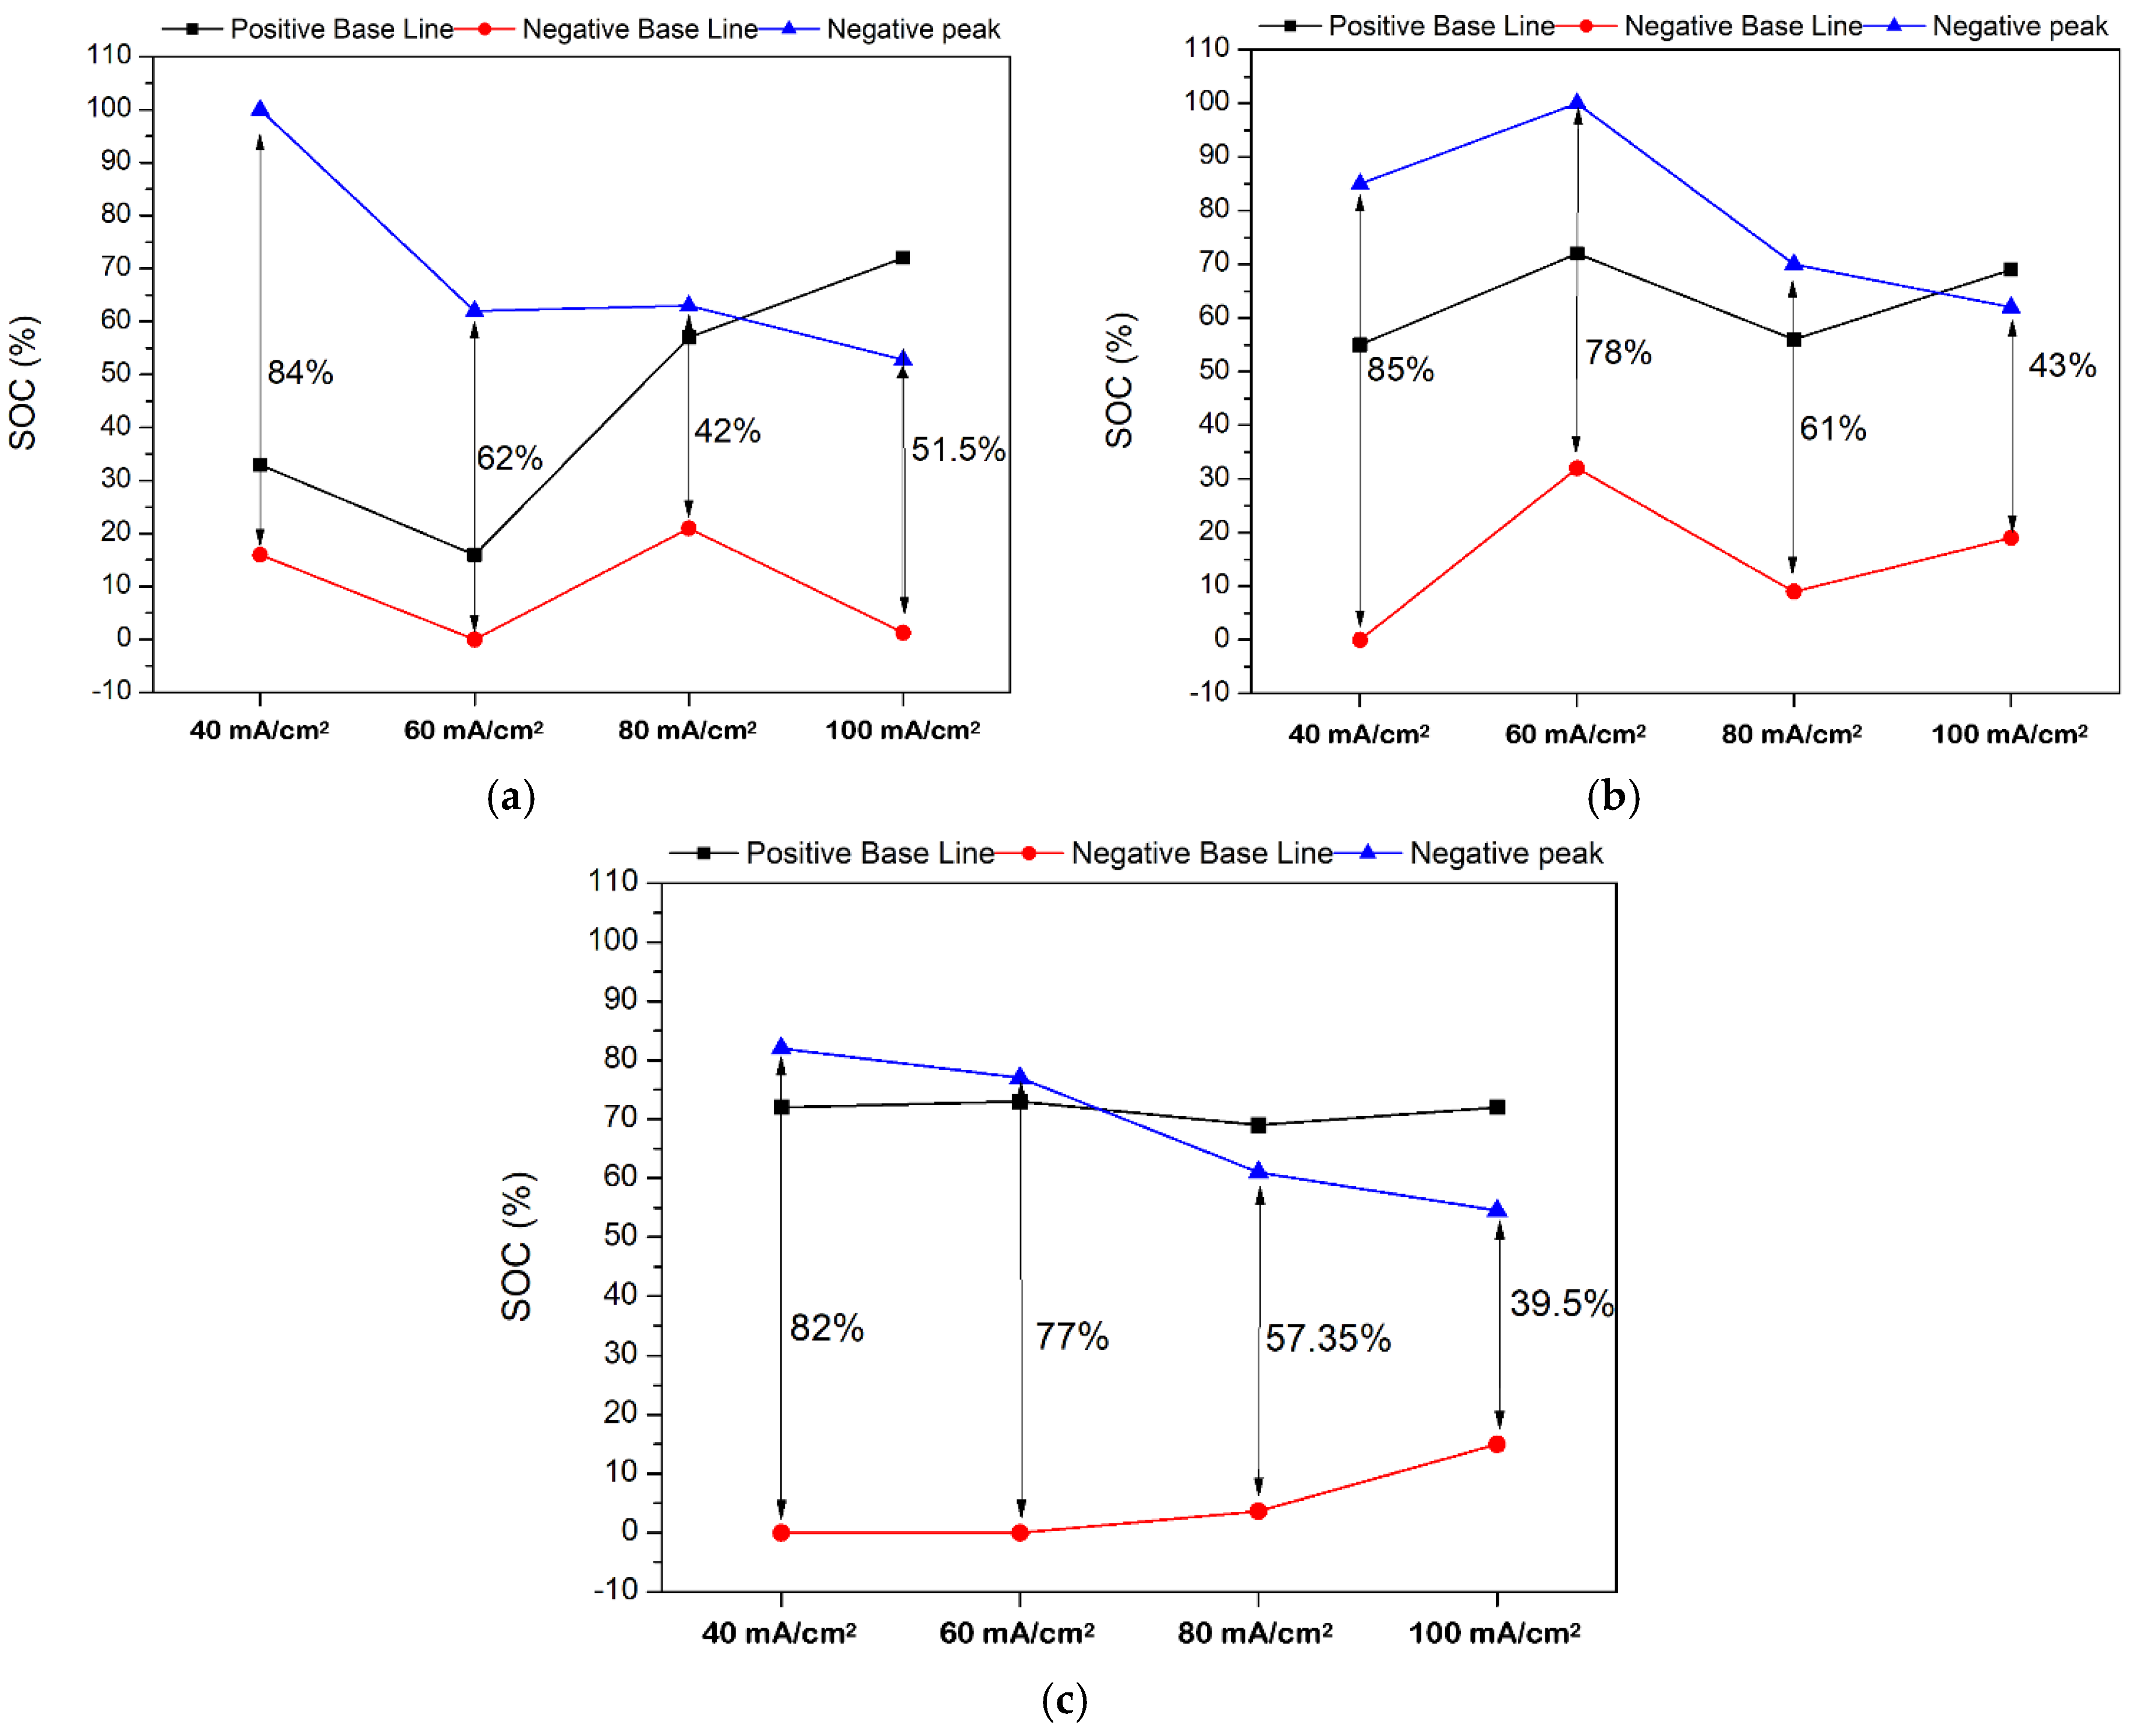
Figure 5 shows the experimental results from the optical sensors monitoring the catholyte (positive electrode) and anolyte (negative electrode) for electrolyte batch #2306003 during 5 charge–discharge cycles at various current densities. Analysis of these results reveals a capacity fading trend as tracked by the optical SOC sensor. Concurrently, the detection signal from the catholyte diminishes with increasing current density, a phenomenon indicating lower electrolyte utilization under high-current-density operation. To investigate the effect of current density on UE, we analyzed the optical SOC range during 5-cycle short-term tests. The results consistently demonstrated that as the current density increased, the electrolyte utilization efficiency significantly decreased. As shown in Figure 6, for batch #2306001, the utilization dropped from 84% at 40 mA/cm2 to 51.5% at 100 mA/cm2. Similarly, utilization for batch #2306002 decreased from 85% to 43%, and for batch #2306003, from 82% to 39.5%. Furthermore, a comparison across batches reveals a trend of decreasing electrolyte utilization with increasing sulfuric acid concentration. This phenomenon is attributed to increased polarization losses at higher current densities, particularly from concentration and activation overpotentials. These losses cause the cell voltage to reach the charge (1.6 V) or discharge (0.7 V) cutoff limits prematurely, thus preventing the completion of the electrochemical reactions. The optical SOC data confirmed this observation: at higher current densities, the peak SOC of the anolyte was markedly lower, indicating that the reduction of V3+ to V2+ was suppressed. Concurrently, for the catholyte, the characteristic absorption peak at high SOC (>90%) diminished or even disappeared with increasing current density, implying that the conversion of active species (oxidation of V4+ to V5+) was also inhibited. The optical SOC monitoring provides direct evidence of the suppressed electrolyte utilization at high current densities.
Figure 5.
For electrolyte batch #2306003, the SOC values for the catholyte and anolyte, as determined by optical sensors and a polynomial conversion model, are shown at various current densities: (a) 40 mA/cm2, (b) 60 mA/cm2, (c) 80 mA/cm2, and (d) 100 mA/cm2, plotted against the cumulative electrical energy.
Figure 6.
Analysis of electrolyte UE for different electrolyte batches under various current densities: (a) #2306001, (b) #2306002, and (c) #2306003.
4.3. Long-Term Cycling Performance, Efficiency, and Capacity Fading
During the 50-cycle galvanostatic aging tests, real-time optical SOC data provided deep insights into the long-term evolution of battery performance.
4.3.1. Capacity Fading and Ion Imbalance Monitored by Optical SOC
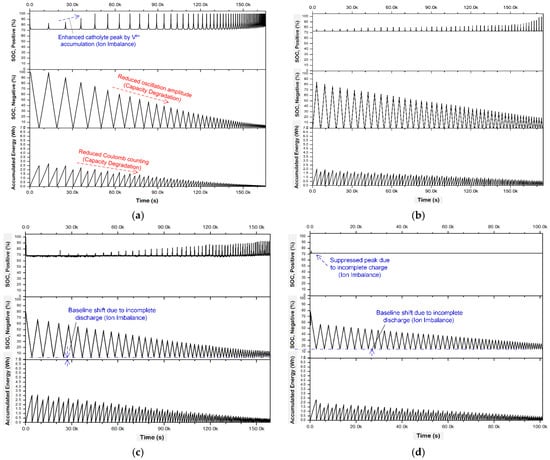
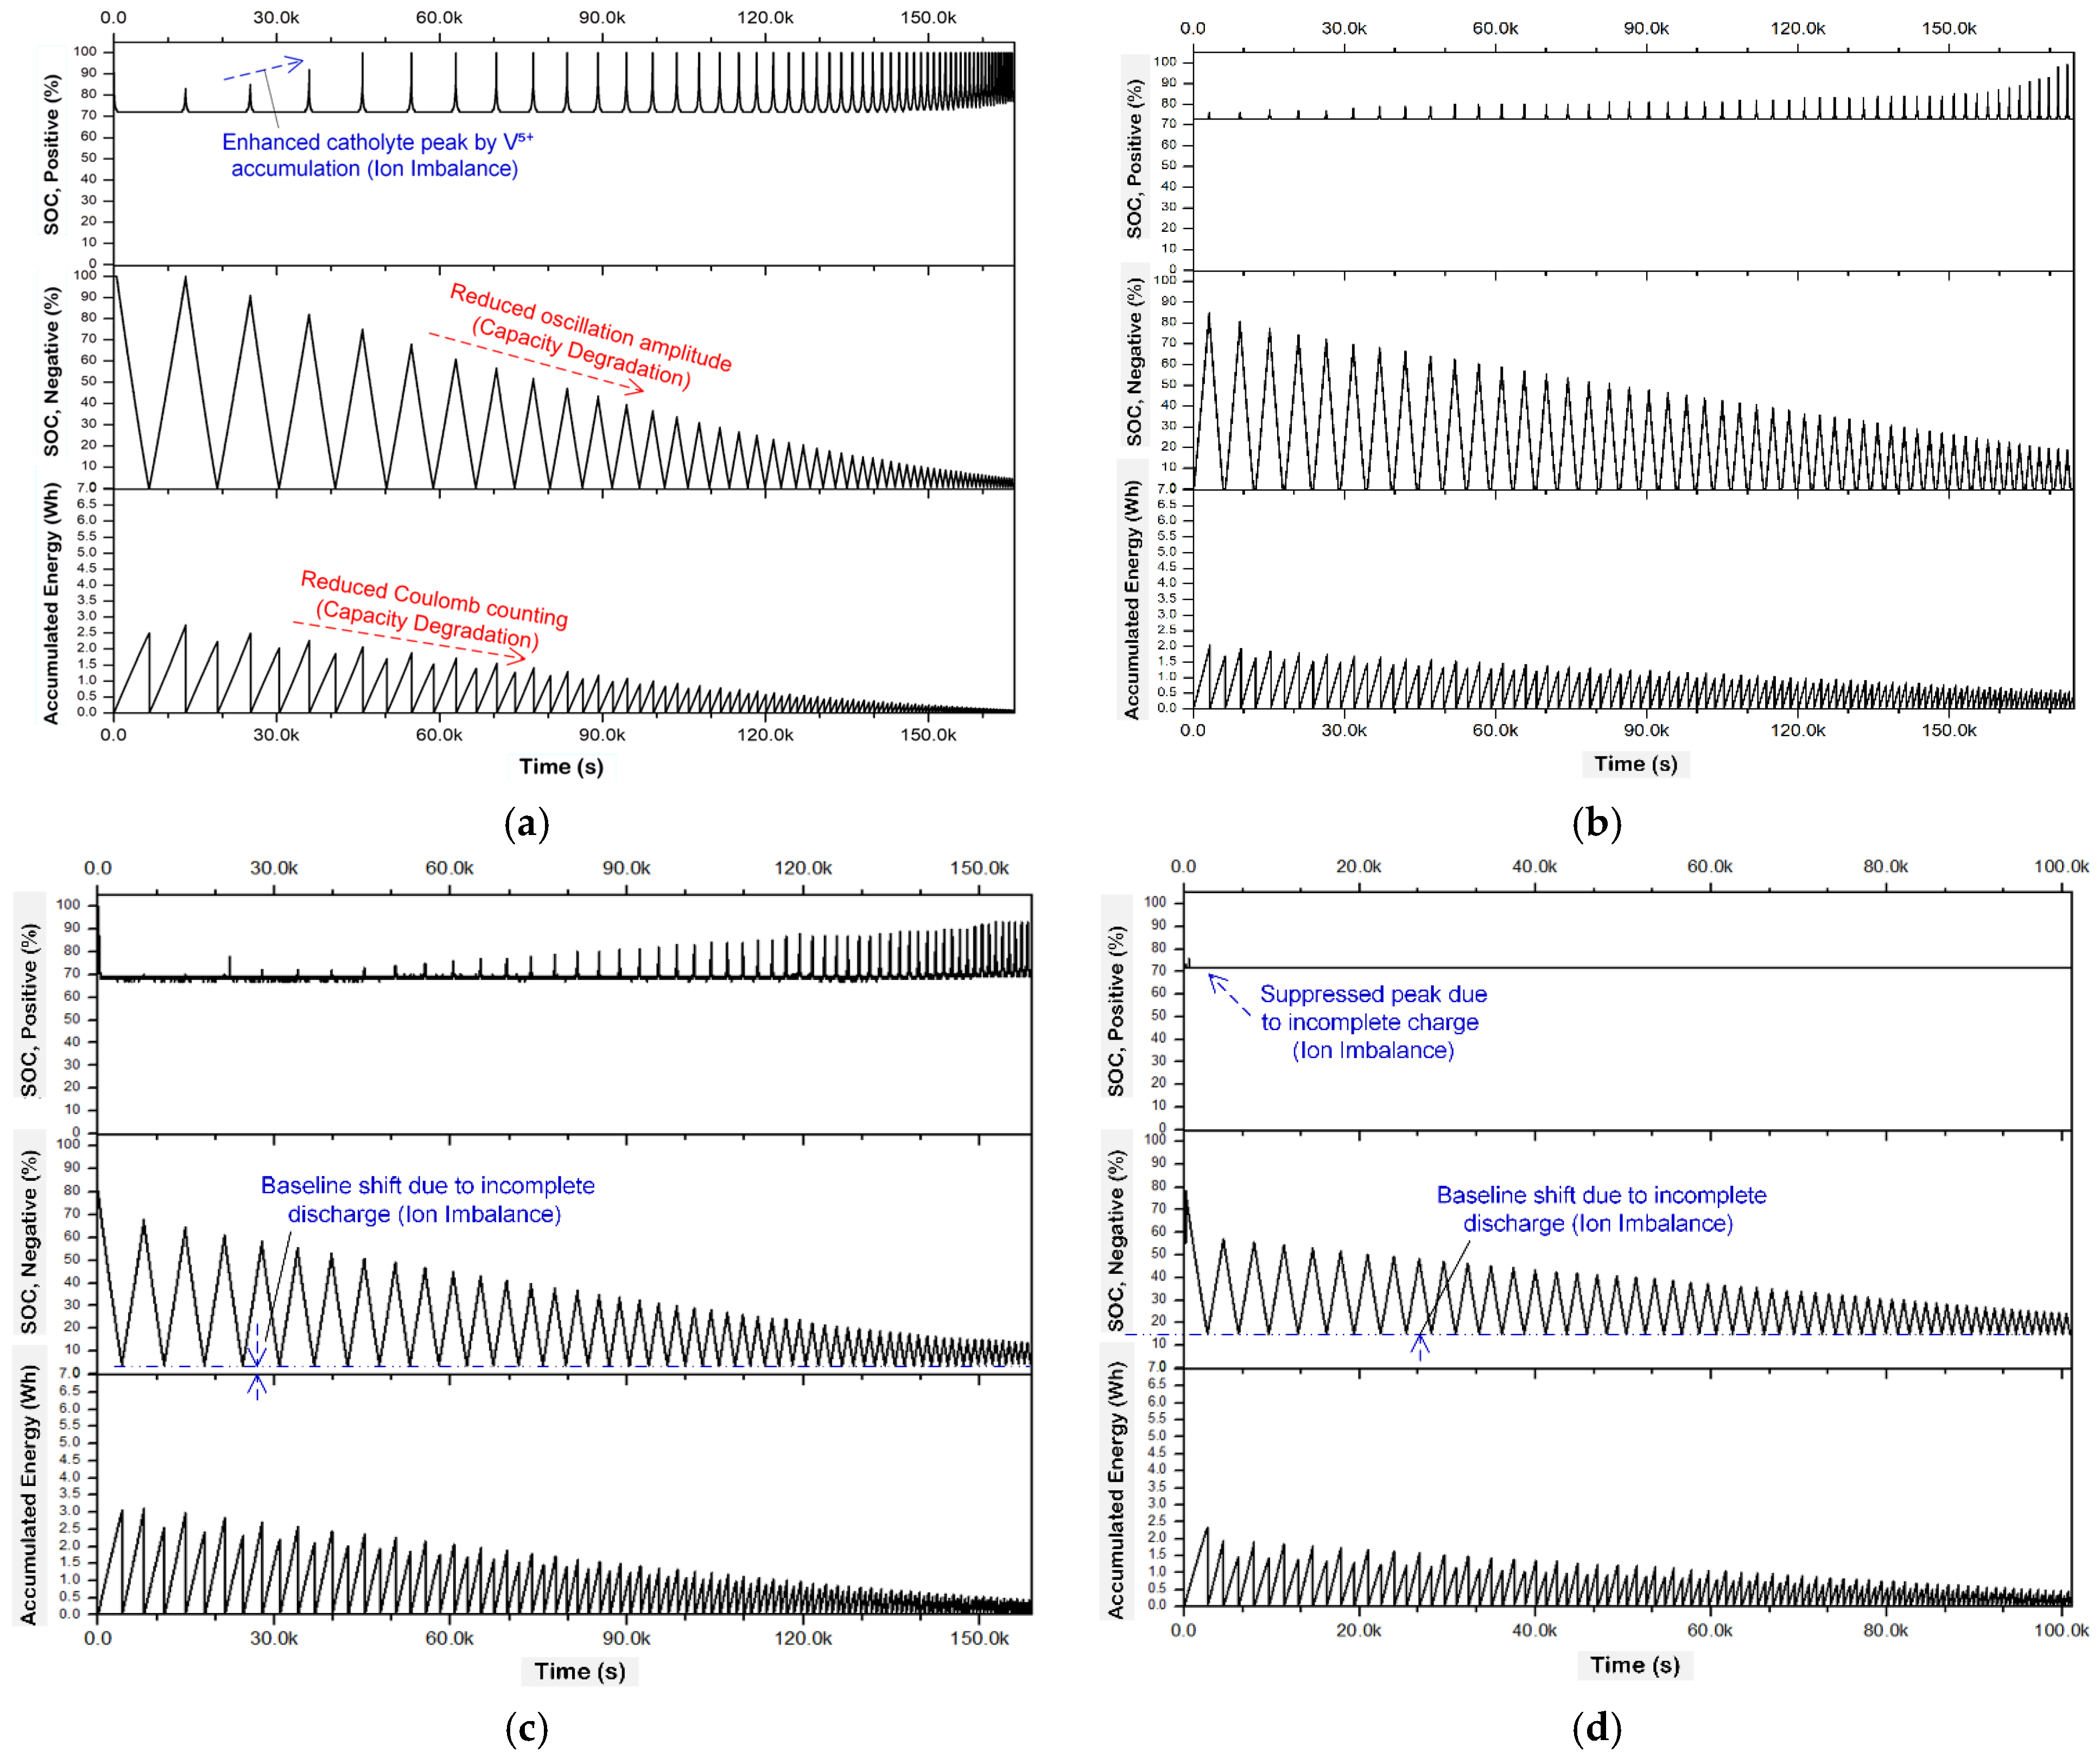
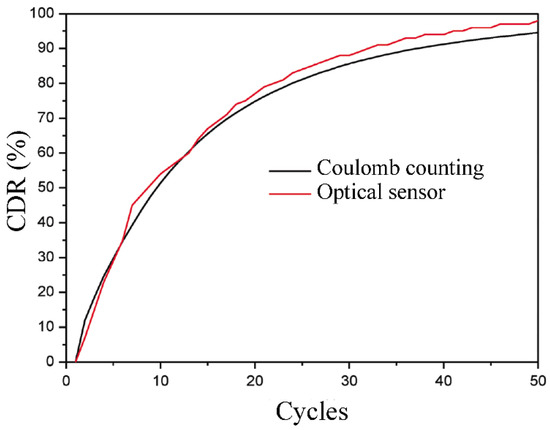
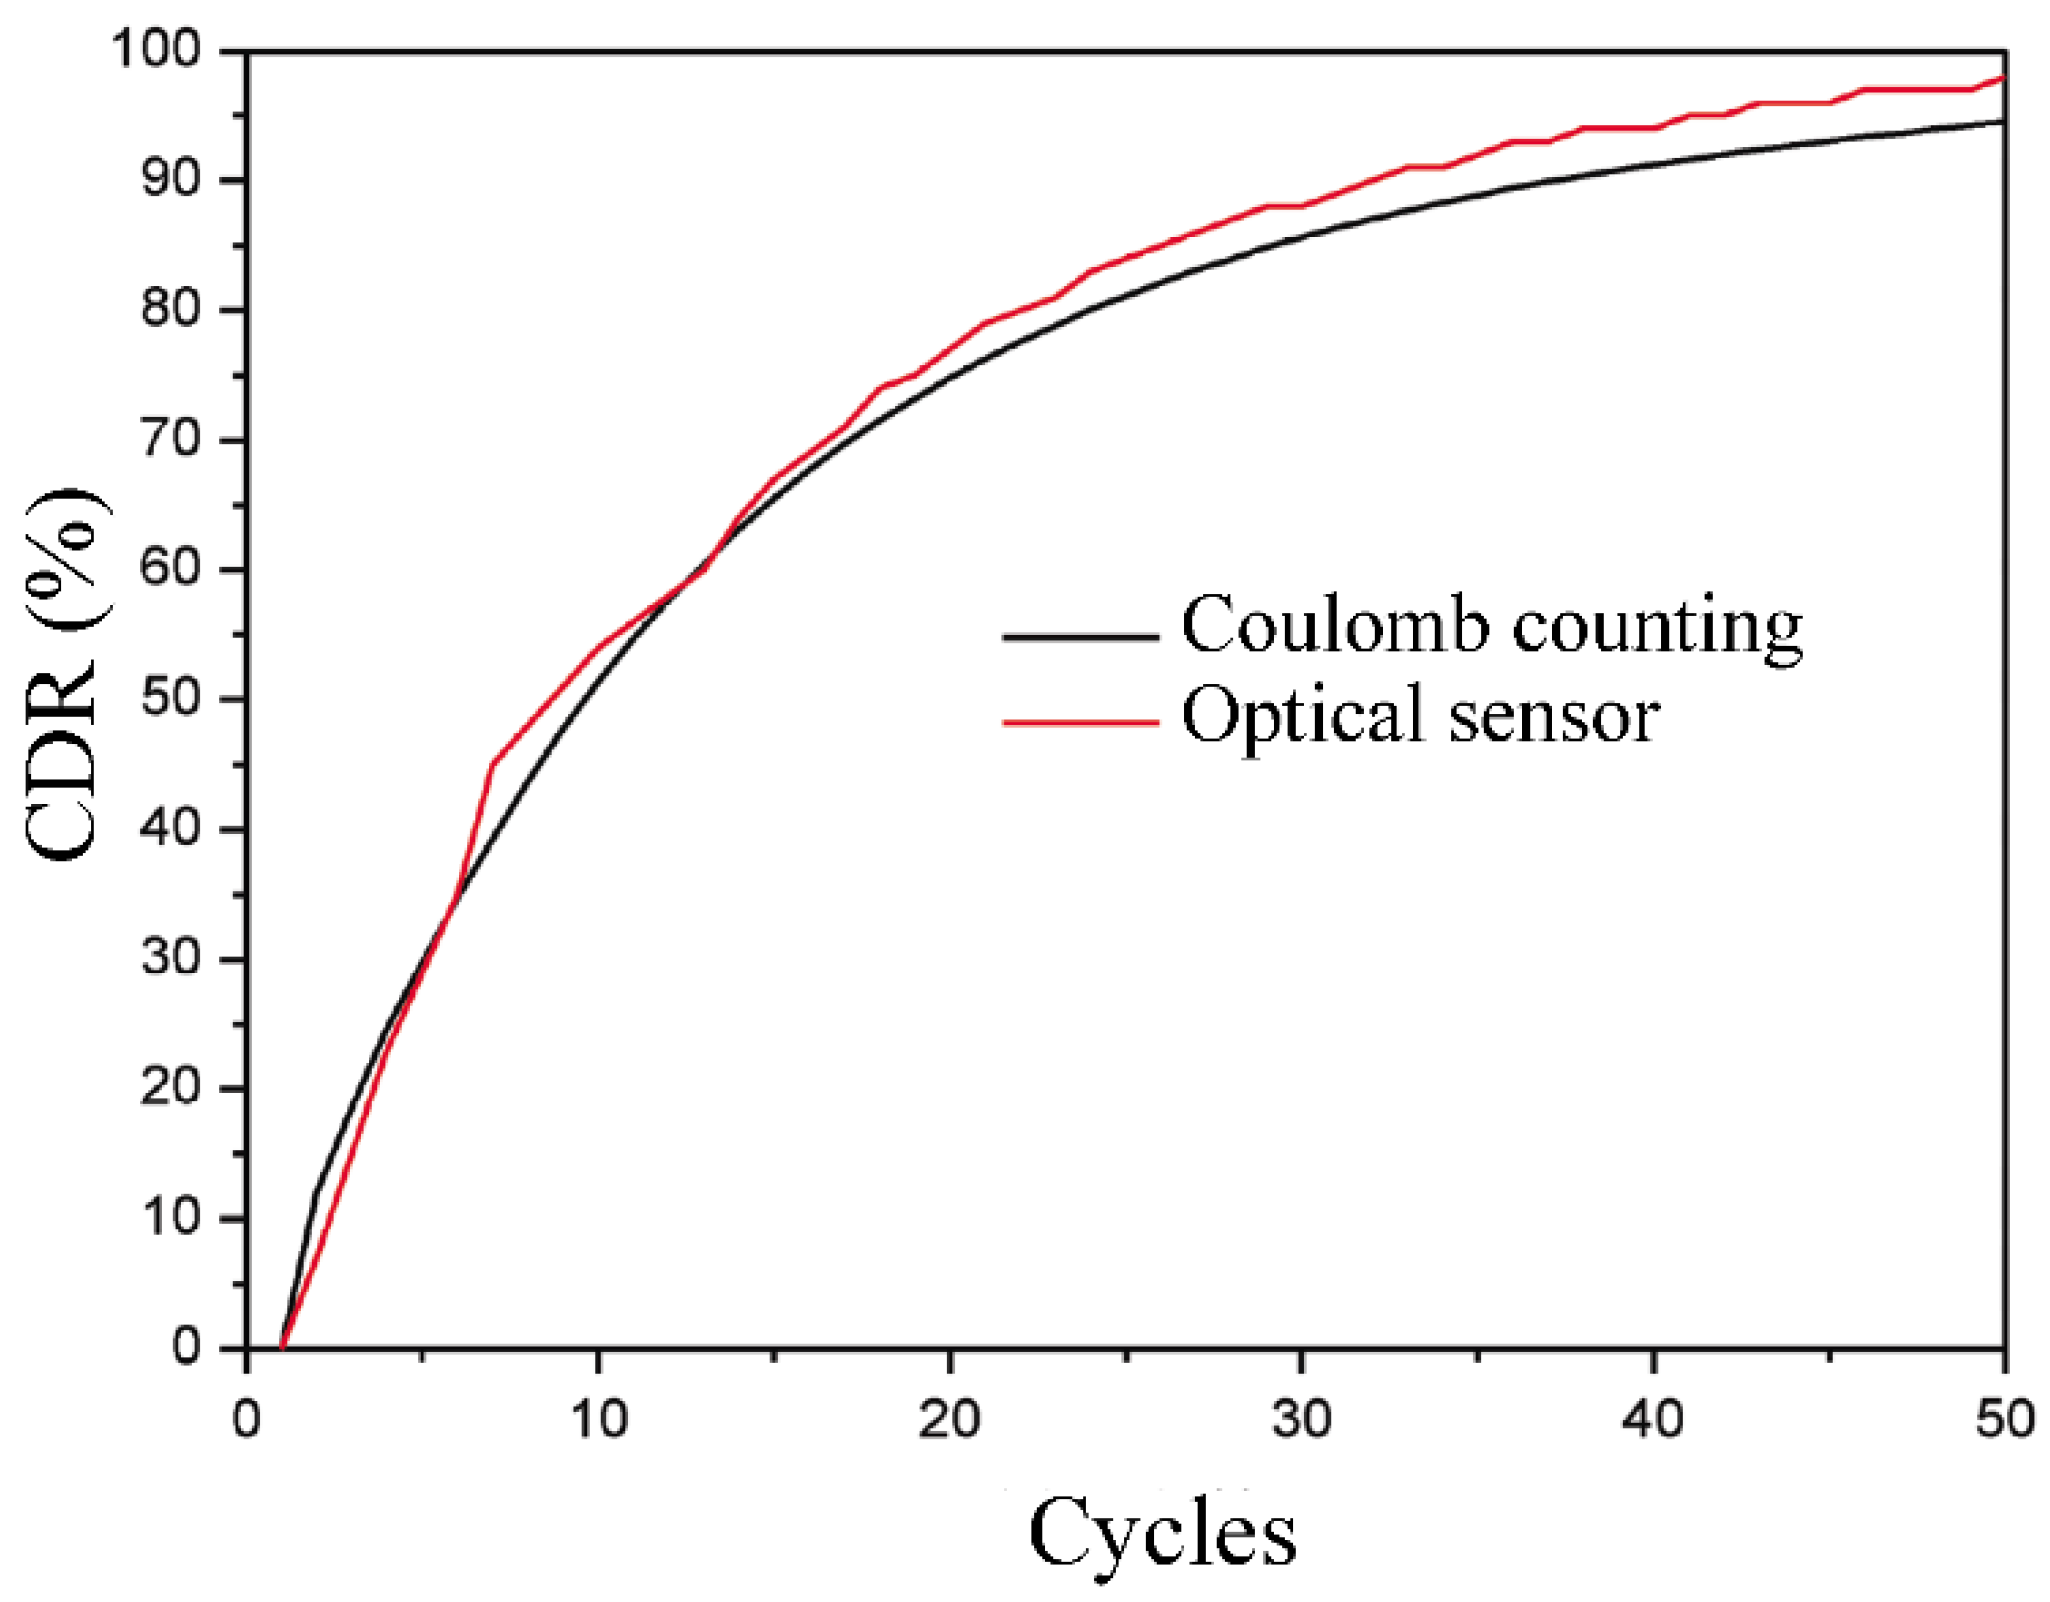
Using the data from electrolyte batch #2306003 as an example, as depicted in Figure 7, the optical SOC data across all test conditions clearly demonstrate a key trend: the amplitude of the anolyte SOC oscillation (i.e., the difference between the peak and trough SOC in each cycle) gradually decreases with an increasing number of cycles, a direct reflection of the battery’s decaying available capacity. As explicitly shown in Figure 8, the capacity degradation rate (CDR) curve derived from the negative electrode optical SOC peaks highly approximates the CDR curve obtained by the battery cycler’s coulomb counting method (exemplified by Batch #2306001 at 40 mA/cm2). This robust synchronous correlation demonstrates that the optical SOC detection technology can accurately resolve the state of the VRFB electrolyte and can be reliably employed for tracking long-term SOH and capacity fading.
Figure 7.
Results of the 50-cycle galvanostatic charge–discharge test for electrolyte batch #2306003 under different constant current densities: (a) 40 mA/cm2, (b) 60 mA/cm2, (c) 80 mA/cm2, and (d) 100 mA/cm2.
Figure 8.
Comparative analysis of capacity degradation rate (CDR) over 50 cycles: optical SOC peak vs. Coulomb counting (batch #2306001 at 40 mA/cm2).
Beyond quantifying the degree of capacity fade, the optical sensors also reveal the direction and signs of ion imbalance. During cycling at low current densities (e.g., 40 mA/cm2), the optical signal from the catholyte at the end of charge (>90% SOC) becomes progressively more pronounced with cycling (Figure 7a). This observation suggests an accumulation of an unintended species. Given the charge direction, this indicates that a greater quantity of V5+ ions remain in the catholyte after each discharge, leading to an apparent increase in the SOC swing on the positive side. Conversely, under high current density conditions (e.g., 100 mA/cm2), as seen in Figure 7d, this peak in the catholyte’s optical SOC is suppressed and may even disappear. This suppression suggests that incomplete charging occurs at high current densities, where polarization effects cause the cell to reach its cutoff voltage prematurely, preventing the full oxidation of V4+ ions into a detectable concentration of V5+ ions. Concurrently, the amount of V2+ generated during each charge cycle progressively decreases. These two observations collectively confirm the phenomenon of an imbalance in active ion concentrations between the catholyte and anolyte (see Figure 7c,d). Furthermore, under high current densities (e.g., 60, 80, and 100 mA/cm2), the data for batch No. #2306003 show that the baseline position of the anolyte optical SOC (at the end of discharge) gradually shifts upward with cycling, consistently failing to return to 0% SOC. This signifies incomplete discharge at high current densities, where a portion of V2+ ions is not fully oxidized back to V3+, thereby further exacerbating the ion imbalance.
4.3.2. Conversion Efficiency and the Effect of Sulfuric Acid Concentration
A summary of the conversion efficiencies, CE, VE, and EE, for the three electrolyte batches before (Initial) and after (Final) 50 cycles is depicted in Table 3. At the initial state, data across most current densities show an energy efficiency ranking of #2306003 > #2306002 > #2306001. This clearly indicates that increasing the sulfuric acid concentration in the electrolyte enhances the initial EE. This effect is likely attributed to the higher electrolyte conductivity associated with increased acid concentration, which reduces the battery’s ohmic losses and subsequently improves VE. Following 50 aging cycles (Final), batches #2306002 and #2306003 generally maintain superior EEs compared to #2306001, demonstrating their better long-term efficiency stability.
Table 3.
Summary of the conversion efficiencies: CE, VE, and EE for the different electrolyte batches before and after 50 cycles.
4.3.3. CDR Analysis
The analysis of CDR reveals the critical impact of operating conditions on battery longevity. As shown in Table 4, a prominent and counter-intuitive trend emerged: for a given electrolyte batch, a higher current density resulted in a lower CDR after 50 cycles. For instance, the CDR for batch #2306001 decreased from 97% at 40 mA/cm2 to 59% at 100 mA/cm2. This counter-intuitive phenomenon is primarily driven by the time-dependent kinetics of parasitic side reactions. The extended charge–discharge periods associated with low current densities maximize the temporal window for these reactions to occur. Specifically, the unstable V(II) species are exposed to oxidation for a longer duration per cycle, thereby accelerating capacity fading. These V(II) ions are highly susceptible to oxidation, such as reaction with residual oxygen or hydrogen evolution. Therefore, operating at high current densities effectively mitigates the long-term capacity degradation rate by drastically reducing the total time the unstable V(II) species are exposed to these capacity-fading mechanisms per Ah discharged. This reduced exposure time translates to poorer CE over extended cycling, which exacerbates the cumulative effects of ion crossover and side reactions (e.g., water transport), thereby accelerating ion imbalance and capacity fade. While time-dependent side reactions are the primary explanation, it is also important to consider the underlying transport limitations. The local heating generated by Ohmic resistance varies significantly with current density, which can induce micro-scale temperature gradients within the flow cell and near the membrane interface. Temperature substantially influences electrolyte properties (e.g., viscosity and conductivity) and ion diffusivity. Under dynamic operation, rapid current changes may induce a lag in ion diffusion and mass transport, particularly at high current densities or during prolonged low-current operation, contributing to the observed utilization suppression and eventual capacity fade. However, when comparing batches with different sulfuric acid concentrations, no clear monotonic relationship was found between capacity retention and acid concentration. As shown in Table 4, at 40 mA/cm2, batch #2306002 showed the lowest degradation; at 60 and 80 mA/cm2, batch #2306003 was superior; and at 100 mA/cm2, batch #2306001 performed best. This complexity highlights that the concentration of the supporting electrolyte, and by extension the proton (H+) concentration, acts as a sophisticated interactive factor impacting long-term capacity maintenance, supporting the necessity to discuss H+ influence even without detailed ion activity validation.
Table 4.
Comparison of capacity degradation rates (CDR %) after 50 cycles.
4.4. UV-Vis Spectroscopic Analysis of the Electrolyte
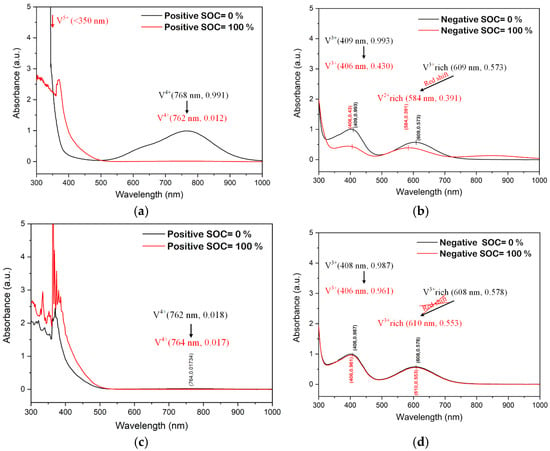
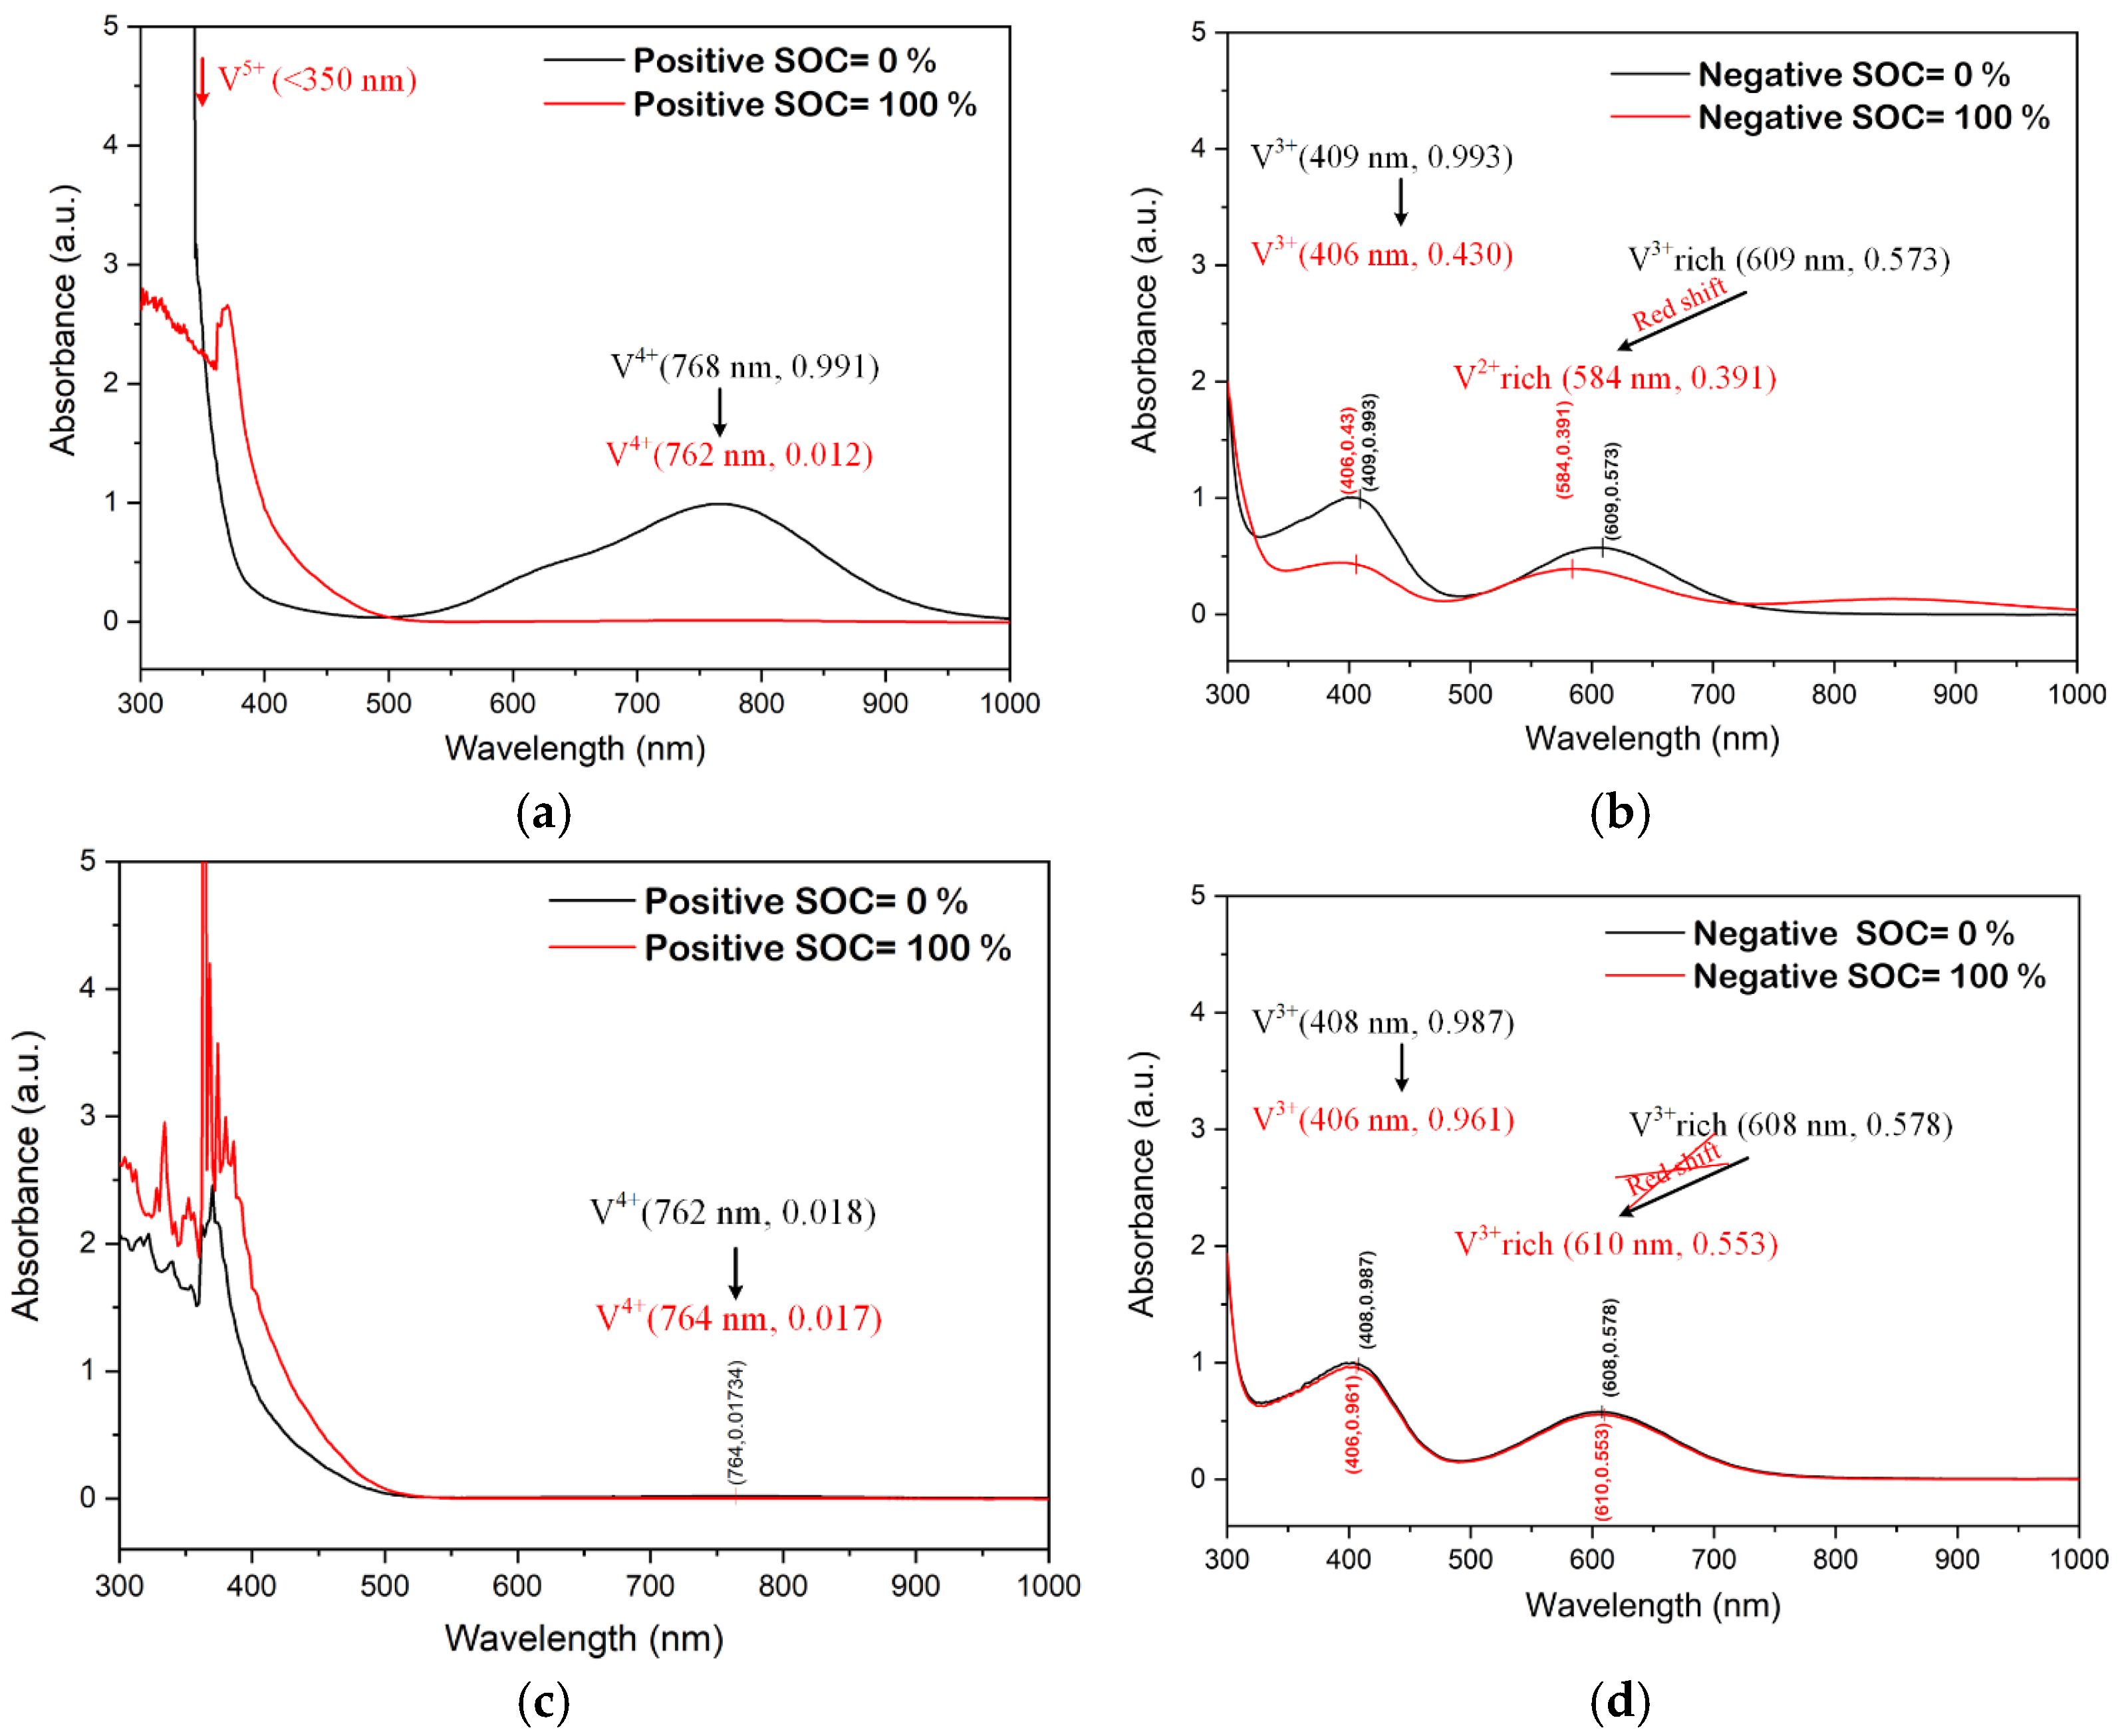
UV-Vis spectroscopic analysis provided more direct, molecular-level evidence of the ion imbalance phenomenon. This study focused on analyzing characteristic peaks at specific wavelengths: the 760 nm peak representing V4+ in the catholyte, the 400 nm peak representing V3+ in the anolyte, and the 570–600 nm band corresponding to the V2+/V3+ transition. The results clearly showed that electrolytes cycled at low current densities (especially 40 mA/cm2) exhibited the largest spectral deviations from their initial states, indicating the most severe degree of ion imbalance. Furthermore, the pronounced variations in optical sensor sensitivity observed across the three batches with differing sulfuric acid concentrations (#2306001: 2.4 M, #2306002: 2.65 M, #2306003: 2.9 M) strongly confirm the critical influence of H+ concentration on the light absorption characteristics of the vanadium species. For instance, from Table 2, batch #2306002 exhibited the optimal optical response (3.51 V voltage span, 0.14% SOC resolution), suggesting that a 2.65 M H2SO4 concentration provided a supporting environment that maximized the concentration-dependent optical signal detected by the sensor. This result emphasizes that precise SOC monitoring requires careful calibration against the specific supporting acid concentration. In Figure 9, using batch #2306001 cycled at 40 mA/cm2 as an example, a comparison of its spectra before and after 50 cycles reveals significant discrepancies. Post-aging spectral analysis revealed severe ion imbalance. At 0% SOC, the diminished intensity of the 760 nm peak indicates incomplete reduction of V5+ in the catholyte. Concurrently, the persistently strong absorbance at 400 nm in the anolyte suggests a significant portion of V3+ failed to convert to V2+. These spectral signatures provide direct evidence of the cumulative capacity loss mechanism (Figure 9d).
Figure 9.
Comparison of UV-Vis spectra for batch 2306001 before and after cycling at 40 mA/cm2: (a) Catholyte initial state, (b) Anolyte initial state, (c) Catholyte aged state, (d) Anolyte aged state.
Based on this analytical approach, the 50-cycle experiments conducted on this batch, utilizing different current densities, were followed by UV-Vis spectroscopic analysis before and after the test, as shown in Table 5. The results of the UV-Vis analysis were quantified to determine the degree of peak change in the electrolyte. For batch #2306001 electrolyte, the ion imbalance severity ranking was determined to be 80 > 40 > 60 > 100 mA/cm2. The imbalance severity rankings for all electrolyte batches, obtained through the same procedure, are collectively summarized in Table 6.
Table 5.
Wavelength and absorbance of characteristic UV-Vis spectral peaks for electrolyte batch #2306001 at fully discharged (SOC = 0%) and fully charged (SOC = 100%) states, before and after 50 charge–discharge cycles.
Table 6.
Ranking of ion imbalance severity after 50 cycles based on UV-Vis spectroscopic analysis.
A strong correlation was found when comparing the degree of spectral imbalance ranked for each current density (Table 5) with the corresponding CDR (Table 4). For batch #2306002, the order of imbalance severity determined by UV-Vis was 40 > 60 > 80 > 100 mA/cm2, which perfectly matches the order of its capacity degradation rates (94.5% > 92.4% > 86.3% > 82.5% at 40, 60, 80, and 100 mA/cm2, respectively). Similarly, for batch #2306003, the imbalance order of 40 > 80 > 100 > 60 mA/cm2 also aligned perfectly with its CDR order (95.9% > 84.4% > 74.2% > 71.8% at 40, 80, 100, and 60 mA/cm2, respectively). This strong correlation suggests that the poor CE at low current densities is the primary driver of ion imbalance, which in turn accelerates capacity fading. The extended charge–discharge times associated with low-current operation allow for greater cumulative ion crossover and side reactions, leading to poorer CE and exacerbating the imbalance.
5. Conclusions
This study has successfully developed and validated a non-invasive optical sensor module for real-time SOC and state-of-health monitoring in VRFBs. By integrating electrochemical performance analysis, in-operando optical monitoring, and UV-Vis spectra characterization, the following main conclusions were drawn:
- The developed optical SOC sensor demonstrates high precision and stability, exhibiting a batch-to-batch resolution between 0.14% and 0.44%, a reproducibility standard deviation of 1.38% to 1.44% in repeated cycling tests, and a long-term static drift rate of 0.509% over 24 h. The sensor can accurately and instantly track the SOC of the negative electrolyte and effectively monitor capacity fading over long-term cycling. The degradation curve derived from the optical SOC peaks shows a trend that highly correlates with the one obtained from the conventional coulomb counting method, demonstrating the feasibility and reliability of this technology for assessing batteries’ state of health.
- Electrolyte composition significantly impacts the optical monitoring capability and battery performance. Increasing the sulfuric acid concentration (from 2.4 M to 2.9 M) effectively enhances the initial EE. Crucially, the optical sensor’s sensitivity (voltage span) and resulting SOC resolution were found to be highly dependent on the electrolyte batch. This pronounced batch variability necessitates batch-specific calibration of the polynomial model, confirming that the model lacks universal generalization capability across different sulfuric acid concentrations. Furthermore, within the concentration range tested, no clear monotonic relationship was observed between sulfuric acid concentration and long-term capacity retention.
- Although operating at higher current densities reduces per-cycle electrolyte utilization due to polarization, it effectively mitigates the long-term capacity degradation rate. This protective effect arises from the minimized chronological exposure of highly reactive V(II) species to parasitic side reactions, thereby preserving the coulombic efficiency over the battery’s lifespan. Conversely, low current density, while achieving high utilization, leads to faster capacity fade because of more pronounced ion imbalance over longer charge–discharge times.
- By combining optical SOC monitoring with UV-Vis spectroscopy, this study provides critical insights into ion imbalance as the core mechanism of capacity fading. The strong consistency between the degree of ion imbalance shown by UV-Vis spectroscopy and the electrochemical performance data (CDR) demonstrates the potential of optical methods for assessing the state of health of the electrolyte.
- Future research should focus on integrating a temperature–current density coupled model to fully capture the complex transport phenomena within VRFBs. Investigating the influence of locally induced thermal gradients on ion diffusivity and crossover kinetics will be essential for developing more precise predictive maintenance models.
- Given that the flow rate was fixed (70 mL/min) across all current density tests, future work should include comparative experiments using varying electrolyte flow rates. Decoupling the effects of current density (reaction rate) from the electrolyte’s residence time (mass transport) within the cell is crucial for fully optimizing operating conditions and validating the time-dependent degradation mechanism.
In summary, this non-invasive optical monitoring technology offers a low-cost and effective solution for the intelligent management of VRFBs. It facilitates precise state awareness and enables proactive maintenance strategies, such as timely electrolyte rebalancing, thereby extending system lifespan and enhancing its operational reliability and economic viability.
Supplementary Materials
The following supporting information can be downloaded at: https://www.mdpi.com/article/10.3390/en18236366/s1, Figure S1: Comparison of polynomial regression models for optical SOC calibration using anolyte data from 5 charge-discharge cycles (Batch #2306003): (a) 5th-order fit and (b) 9th-order fit. Both models demonstrate a high goodness of fit with R2 = 0.99997. Table S1: Reproducibility test results and estimated optical sensor error for batch #2306001, conducted across 15 conditions (3 current densities and 5 electrolyte flow rates) with 3 replicates over 5 cycles.
Author Contributions
Conceptualization, Y.-M.W.; Methodology, Y.-M.W. and S.-C.C.; Validation, S.-C.C.; Formal Analysis, C.-C.C.; Investigation, S.-C.C.; Resources, C.-H.K. and H.-J.L.; Writing—Original Draft Preparation, C.-C.C. and S.-C.C.; Writing—Review & Editing, C.-C.C., Y.-M.W. and N.-Y.H.; Visualization, S.-C.C.; Supervision, C.-H.K., H.-J.L. and J.-Y.K.; Project Administration, J.-Y.K.; Funding Acquisition, C.-C.C., Y.-M.W. and N.-Y.H. All authors have read and agreed to the published version of the manuscript.
Funding
This research was funded by the National Atomic Research Institute (NARI) of Taiwan, grant number 114-C1005, and the National Science and Technology Council (NSTC) of Taiwan, grant numbers NSTC 112-2622-E-019-005 and NSTC 113-2622-E-019-006.
Data Availability Statement
The original contributions presented in the study are included in the article. Further inquiries can be directed to the corresponding author.
Acknowledgments
The authors gratefully acknowledge the valuable technical support and assistance provided by Sheng-Yu Wang and Ming-Tai Su from the Metal Industries Research & Development Centre, and by Jui-Hsiung Huang and Hong-Yu Lin from the Green Technology Research Institute (GTRI) of CPC Corporation.
Conflicts of Interest
Authors Shang-Ching Chuang and Yao-Ming Wang were employed by the Metal Industries Research and Development Centre. The remaining authors declare that the research was conducted in the absence of any commercial or financial relationships that could be construed as a potential conflict of interest.
Abbreviations
The following abbreviations are used in this manuscript:
| VRFB | All-vanadium redox flow battery |
| SOC | State of charge |
| SOH | State of health |
| UV-Vis | Ultraviolet–visible |
| LED | Light-emitting diode |
| CC | Constant current |
| CV | Constant voltage |
| CE | Coulombic efficiency |
| VE | Voltage efficiency |
| EE | Energy efficiency |
| UE | Utilization efficiency |
| CDR | Capacity degradation rate |
References
- Schofield, K.; Musilek, P. State of Charge and Capacity Tracking in Vanadium Redox Flow Battery Systems. Clean Technol. 2022, 4, 607–618. [Google Scholar] [CrossRef] [Scilit]
- Ma, C.-T. A Novel State of Charge Estimating Scheme Based on an Air-Gap Fiber Interferometer Sensor for the Vanadium Redox Flow Battery. Energies 2020, 13, 291. [Google Scholar] [CrossRef] [Scilit]
- Skyllas-Kazacos, M.; Rychcik, M.; Robins, R.G.; Fane, A.G.; Green, M.A. New All-Vanadium Redox Flow Cell. J. Electrochem. Soc. 1986, 133, 1057–1058. [Google Scholar] [CrossRef] [Scilit]
- Sum, E.; Skyllas-Kazacos, M. A study of the V(II)/V(III) redox couple for redox flow cell applications. J. Power Sources 1985, 15, 179–190. [Google Scholar] [CrossRef] [Scilit]
- Sum, E.; Rychcik, M.; Skyllas-kazacos, M. Investigation of the V(V)/V(IV) system for use in the positive half-cell of a redox battery. J. Power Sources 1985, 16, 85–95. [Google Scholar] [CrossRef] [Scilit]
- Geiser, J.; Natter, H.; Hempelmann, R.; Morgenstern, B.; Hegetschweiler, K. Photometrical Determination of the State-of-Charge in Vanadium Redox Flow Batteries Part I: In Combination with Potentiometric Titration. Z. Phys. Chem. 2019, 233, 1683–1694. [Google Scholar] [CrossRef] [Scilit]
- Chou, Y.-S.; Hsu, N.-Y.; Jeng, K.-T.; Chen, K.-H.; Yen, S.-C. A novel ultrasonic velocity sensing approach to monitoring state of charge of vanadium redox flow battery. Appl. Energy 2016, 182, 253–259. [Google Scholar] [CrossRef] [Scilit]
- Feng, R.; Guo, Z.; Meng, X.; Sun, C. Modeling and State of Charge Estimation of Vanadium Redox Flow Batteries: A Review. Energies 2025, 18, 4666. [Google Scholar] [CrossRef] [Scilit]
- Ghimire, P.C.; Bhattarai, A.; Lim, T.M.; Wai, N.; Skyllas-Kazacos, M.; Yan, Q. In-Situ Tools Used in Vanadium Redox Flow Battery Research—Review. Batteries 2021, 7, 53. [Google Scholar] [CrossRef] [Scilit]
- Skyllas-Kazacos, M.; Kazacos, M. State of charge monitoring methods for vanadium redox flow battery control. J. Power Sources 2011, 196, 8822–8827. [Google Scholar] [CrossRef] [Scilit]
- Liu, L.; Xi, J.; Wu, Z.; Zhang, W.; Zhou, H.; Li, W.; Qiu, X. State of charge monitoring for vanadium redox flow batteries by the transmission spectra of V(IV)/V(V) electrolytes. J. Appl. Electrochem. 2012, 42, 1025–1031. [Google Scholar] [CrossRef] [Scilit]
- Tang, Z.; Aaron, D.S.; Papandrew, A.B.; Zawodzinski, T.A. Monitoring the State of Charge of Operating Vanadium Redox Flow Batteries. ECS Trans. 2012, 41, 1. [Google Scholar] [CrossRef] [Scilit]
- Loktionov, P.; Pichugov, R.; Konev, D.; Petrov, M.; Pustovalova, A.; Antipov, A. Operando UV/Vis spectra deconvolution for comprehensive electrolytes analysis of vanadium redox flow battery. J. Electroanal. Chem. 2022, 925, 116912. [Google Scholar] [CrossRef] [Scilit]
- Geiser, J.; Natter, H.; Hempelmann, R.; Morgenstern, B.; Hegetschweiler, K. Photometrical Determination of the State-of-Charge in Vanadium Redox Flow Batteries Part II: In Combination with Open-Circuit-Voltage. Z. Phys. Chem. 2019, 233, 1695–1711. [Google Scholar] [CrossRef] [Scilit]
- Zhang, W.; Liu, L.; Liu, L. An on-line spectroscopic monitoring system for the electrolytes in vanadium redox flow batteries. RSC Adv. 2015, 5, 100235–100243. [Google Scholar] [CrossRef] [Scilit]
- Rudolph, S.; Schröder, U.; Bayanov, I.M.; Blenke, K.; Hage, D. High resolution state of charge monitoring of vanadium electrolytes with IR optical sensor. J. Electroanal. Chem. 2013, 694, 17–22. [Google Scholar] [CrossRef] [Scilit]
- Quill, N.; Petchsingh, C.; Lynch, R.P.; Gao, X.; Oboroceanu, D.; Ní Eidhin, D.; O’Mahony, M.; Lenihan, C.; Buckley, D.N. Factors Affecting Spectroscopic State-of-Charge Measurements of Positive and Negative Electrolytes in Vanadium Redox Flow Batteries. ECS Trans. 2015, 64, 23. [Google Scholar] [CrossRef] [Scilit]
- Shin, K.-H.; Jin, C.-S.; So, J.-Y.; Park, S.-K.; Kim, D.-H.; Yeon, S.-H. Real-time monitoring of the state of charge (SOC) in vanadium redox-flow batteries using UV–Vis spectroscopy in operando mode. J. Energy Storage 2020, 27, 101066. [Google Scholar] [CrossRef] [Scilit]

Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2025 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license (https://creativecommons.org/licenses/by/4.0/).