
Article

Copper Coordination Compounds as Corrosion-Resistant Materials for Seawater Electrolysis

1
Centro de Desarrollo Energético Antofagasta, Universidad de Antofagasta, Antofagasta 1240000, Chile
2
Departamento de Química, Facultad de Ciencias Básicas, Universidad de Antofagasta, Antofagasta 1240000, Chile
3
Departamento de Ingeniería Química y Procesos de Minerales, Universidad de Antofagasta, Avenida Universidad de Antofagasta 02800, Antofagasta 1271155, Chile
4
Departamento de Ingeniería en Metalurgia, Universidad de Atacama, Av. Copayapu 485, Copiapó 1530000, Chile
5
Departamento de Química de los Materiales, Facultad de Química y Biología, Universidad de Santiago de Chile, Av. L. B. O’Higgins 3363, Casilla 40, Correo 33, Santiago 3840000, Chile
6
Departamento de Química y Biología, Facultad de Ciencias Naturales, Universidad de Atacama, Av. Copayapú 485, Copiapó 1530000, Chile
7
Faculty of Engineering and Architecture, Universidad Arturo Prat, Avenida Arturo Prat 2120, Iquique 1110939, Chile
*
Author to whom correspondence should be addressed.
Processes 2026, 14(3), 423; https://doi.org/10.3390/pr14030423
Submission received: 7 January 2026 / Revised: 20 January 2026 / Accepted: 22 January 2026 / Published: 25 January 2026

Abstract

Seawater electrolysis offers a promising route for sustainable hydrogen production in coastal areas, leveraging solar energy while reducing freshwater consumption. Yet, chloride-induced corrosion severely limits conventional electrodes such as titanium, which depend on passive titanium dioxide films and display minimal hydrogen evolution reaction activity (|i0,H2| ≈ 0.001–0.01 A/m2). Here, we report for the first time the use of copper-based coordination compounds—a triazole-derived polymer (CCCu) and a Prussian Blue Analogue (CuHCF)—as dual-function electrodes combining corrosion resistance with electrocatalytic activity. Structural integrity was verified by FTIR, TGA, XRD, and SEM/EDS analyses. Electrochemical tests in 0.5 M NaCl, interpreted using mixed potential theory, revealed corrosion potentials (Ecorr) of −40 mV versus Standard Hydrogen Electrode (CuHCF) and −23 mV versus Standard Hydrogen Electrode (CCCu), and corrosion current densities of 0.259 and 0.379 A/m2, respectively. Both exhibited hydrogen evolution reaction exchange current densities significantly higher than titanium (0.019 A/m2 for CuHCF and 0.062 A/m2 for CCCu). CuHCF achieved a Tafel slope of 222 mV/dec, comparable to NiMoP alloys and carbon steel. Complementary density functional theory calculations elucidated how metal–ligand interactions and electronic redistribution govern both catalytic performance and degradation. These findings introduce a new concept of semi-electrocatalysts, where copper coordination compounds act as structurally adaptive, low-cost materials bridging corrosion resistance and hydrogen evolution in seawater systems.

1. Introduction

Seawater electrolysis has emerged as a promising and sustainable alternative for hydrogen (H2) production in coastal regions with abundant solar resources, such as northern Chile (Atacama Desert), due to the vast availability of both seawater and sunlight. Among renewable energy sources, solar energy is the most widely utilized, giving rise to the concept of solar hydrogen. This process offers significant environmental and operational advantages, including high energy-conversion efficiency and the production of high-purity hydrogen, with chlorine (Cl2) as the only undesired by-product [1,2,3], as illustrated in Figure 1.
Figure 1 presents a conceptual scheme of solar electrolysis in a chloride-containing aqueous medium, using copper hexacyanoferrate (CuHCF) for the cathode electrode and other materials such as RuO2 or IrO2, as the anode [3]. At the cathode, CuHCF facilitates the hydrogen evolution reaction (HER), where water molecules are reduced to generate H2. At the anode, chloride ions undergo oxidation via the chlorine evolution reaction (CER), which competes with the oxygen evolution reaction (OER). A photovoltaic panel supplies the energy required to drive both processes. The scheme illustrates the role of CuHCF as an electrocatalyst enabling the simultaneous production of H2 and Cl2 from saline solutions. Nevertheless, seawater electrolysis remains technically challenging due to the corrosive nature of chloride-rich environments, which accelerate electrode degradation and favor CER over OER [4,5]. Within this context, Prussian Blue Analogues (PBAs), a family of metal–cyanide coordination compounds, have attracted attention as electrode materials for saline electrolysis because of their structural robustness and redox versatility. For OER, PBAs such as cobalt hexacyanoferrate (CoHCF) have been reported as promising anodic materials capable of promoting oxygen evolution while mitigating the detrimental effects associated with chloride-containing electrolytes [6,7,8].
This not only reduces the efficiency of water splitting but also poses significant environmental and safety risks. To mitigate this issue, recent studies have explored materials such as NiFe layered double hydroxides, cobalt phosphides (CoP), and transition-metal oxides like RuO2 and IrO2, which promote selective OER over CER at the anode [9,10,11,12,13].
In this context, the development of materials that combine corrosion resistance with high electrocatalytic activity becomes essential. Coordination polymers (CPs) have attracted increasing attention as multifunctional materials capable of simultaneously addressing both requirements [1,2,14,15,16]. Their ordered structures, composed of metal centers coordinated to organic ligands, allow the rational design of materials with favorable electronic configurations, high porosity, and strong resistance to corrosion in aggressive environments [17,18]. The strength and directionality of coordinate covalent bonds formed through Lewis acid–base interactions between metal ions and donor atoms such as nitrogen or oxygen confer chemical robustness and stability to CPs, even under highly saline and oxidative conditions [19]. By carefully selecting ligands such as triazoles or carboxylates, and metal centers such as Cu2+, Co2+, or Fe3+, CPs can be designed to exhibit high catalytic activity for the HER and oxygen reduction reaction (ORR) at the cathode, while maintaining structural integrity and oxidative resistance at the anode [5,14].
In this dual role, coordination compounds act both as active catalysts and as protective frameworks, mitigating electrochemical degradation while facilitating target reactions [2,9,20,21]. PBAs, a subclass of coordination compounds formed by cyanide-bridged metal networks, offer additional advantages: they are low-cost, easy to synthesize, and feature high surface area, uniform catalytic sites, and excellent tolerance to chloride ions under both anodic and cathodic conditions. Nickel hexacyanoferrate (NiHCF) has demonstrated promising HER performance in alkaline media, achieving current densities of 10 mA/cm2 at an overpotential of just 200 mV [21,22,23,24]. These findings highlight the potential of coordination chemistry in developing electrode materials that promote OER, suppress chlorine generation, and withstand corrosive degradation, thus advancing sustainable hydrogen production from seawater. HER in seawater is more challenging than in pure water or acidic solutions because of the low proton concentration, pH fluctuations, and the deposition of impurities (e.g., Mg2+, Ca2+, heavy metals) on catalyst surfaces, which block active sites and reduce durability. Therefore, strategies such as membranes, corrosion-resistant catalysts, and permselective overlayers are needed to enhance HER performance in seawater [25].
Alternatively, the integration of Density Functional Theory (DFT) calculations provides a valuable complement to the experimental findings, as they make it possible to describe phenomena that cannot be directly measured with current technologies or whose experimental determination would be excessively complex. Within this framework, DFT allows the exploration of reaction pathways, charge redistribution, and the role of metal–ligand interactions in promoting the HER under seawater electrolysis conditions, where multiple competing processes occur and transient intermediates are difficult to track experimentally. On this basis, we propose a reaction mechanism for hydrogen formation supported by experimental data, thereby offering a coherent interpretation that connects structural and electronic features with the observed electrocatalytic performance.
Despite these advances, there are no previous reports on the use of copper-based coordination compounds as bifunctional electrode materials for saline water electrolysis. In this study, we introduce two copper coordination systems: a triazole-derived polymer (CCCu) and a Prussian Blue analogue (CuHCF), highlighting their dual role as corrosion-resistant materials during the HER. To the best of our knowledge, this is the first work that systematically evaluates the electrochemical behavior of these compounds in chloride-rich environments, combining conventional characterization techniques (FTIR, TGA, XRD, SEM/EDS) with a kinetic superposition model based on the mixed potential theory. This integrated approach not only reveals their intrinsic catalytic and stability features but also establishes a new methodological framework for assessing coordination chemistry materials under realistic saline conditions relevant to solar-driven seawater electrolysis.
Beyond its specific contribution to materials science, this research is framed within a broader context in which technological innovation and the efficient use of resources exert transformative impacts on energy systems and water management. Recent studies demonstrate that the integration of renewable energies with sustainable hydrogen production processes generates direct socio-economic benefits, fostering local value chains, promoting the creation of skilled employment, and strengthening human resilience in the face of water scarcity and the effects of climate change. The reuse of saline effluents and the development of corrosion-resistant electrodes in chloride-rich media adhere to the principles of the circular economy, reducing waste and exploiting underutilized resources while expanding the resource base available for the energy transition. In this context, the use of copper coordination compounds advances sustainability, innovation, and equity in the distribution of water and energy resources, thereby contributing tangibly to economic growth and global social well-being [26,27,28,29].
This manuscript reports, for the first time, the application of copper-based coordination compounds as unconventional electrode materials for solar-driven seawater electrolysis. Unlike conventional metals such as titanium (Ti), which rely on passive oxide layers and exhibit negligible catalytic activity, these compounds introduce a new paradigm by combining intrinsic corrosion resistance with active electrocatalytic behavior. Their performance in 0.5 M NaCl is systematically evaluated using a mathematical superposition model based on mixed potential theory, enabling the deconvolution of overlapping electrochemical processes and offering unprecedented insight into their dual functionality. Remarkably, the results reveal that these coordination frameworks not only act as protective matrices but also operate as active “semi-electrocatalysts”, sustaining hydrogen evolution while undergoing controlled structural transformations. This novel approach expands the application field of coordination chemistry, traditionally confined to energy storage and molecular catalysis, toward the design of low-cost, multifunctional electrodes for saline conditions. However, the manuscript does not include long-term stability tests such as chronoamperometry or multi-cycle voltammetry to quantify the operational lifetime of CCCu and CuHCF, since our intention is to study the corrosion effect of these materials when in contact with saline electrolytes during electrolysis.

2. Materials and Methods

2.1. Synthesis of Coordination Compounds

2.1.1. Coordination Compounds of Copper (CCCu)

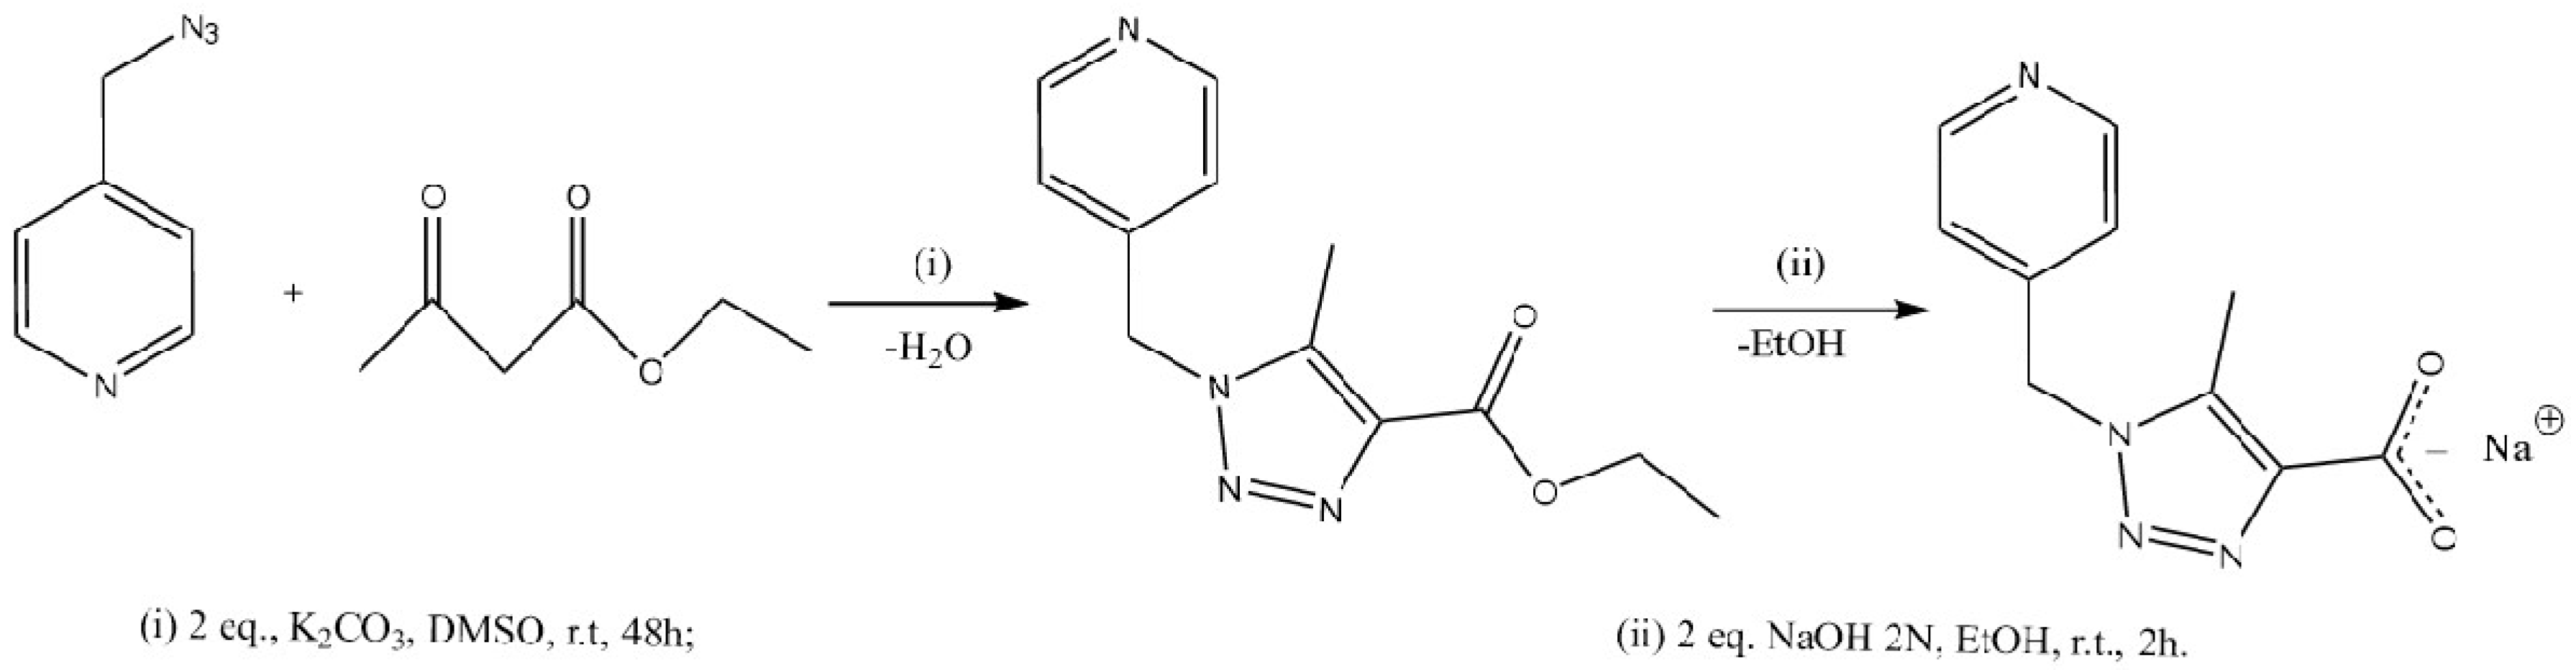
The precursor of the organic ligand, referred to as L1, was synthesized following standard procedures reported in the literature [20], yielding ethyl 5-methyl-1-(pyridin-3-ylmethyl)-1H-1,2,3-triazole-4-carboxylate. The ester was subsequently saponified using a NaOH solution (2 equivalents) in a minimal volume of methanol, resulting in the sodium carboxylate salt (L1). Figure 2 shows a two-step organic synthesis scheme starting from an azide compound and a β-ketoester to produce a triazole derivative substituted with a pyridyl group.
An aqueous solution of L1 (5 mL, 5 mg, 0.230 mmol) was then added dropwise to a solution of copper(II) nitrate (0.195 mmol) in formamide (5 mL) and distilled water (2.5 mL), following an established protocol (Sigma Aldrich Chemical Reagent Co., 3050 Spruce Street, St. Louis, MO, USA), as illustrated in Figure 2. The mixture was maintained at room temperature for two weeks, after which crystals suitable for single-crystal X-ray diffraction (SC-XRD) analysis formed at the interface [30].

2.1.2. CuHCF Synthesis

The CuHCF was synthesized by controlled coprecipitation, following methodologies described in the literature [31,32]. Under stirring, two solutions were prepared: (A) 25 mL of 0.03 M K3[Fe(CN)6] and (B) 75 mL of 0.09 M Cu(NO3)2 (Sigma Aldrich Chemical Reagent Co.). Solution A was added dropwise to solution B (60 °C, 600–700 rpm) to promote uniform nucleation and controlled solid growth. After completing the mixing (final volume 100 mL), stirring was maintained for 4 h at 60 °C. The resulting precipitate was vacuum filtered, washed sequentially with distilled water to remove soluble salts and with ethanol to displace residual water, then dried in an oven at 70 °C for 24 h. Finally, the solid was allowed to cool to room temperature and stored for subsequent analysis [33].

2.2. Characterization

2.2.1. Infrared Spectroscopy (FTIR)

IR spectra were recorded to identify the functional groups and confirm the coordination between the metal centers and the organic ligands in the synthesized compounds. The measurements were performed using a Nicolet Avatar 330 FTIR spectrometer (Thermo Electron Corporation, 5225 Verona Road, Madison, WI, USA), equipped with a DTGS (Deuterated Triglycine Sulfate) detector and a KBr beam splitter for enhanced sensitivity and resolution in the mid-infrared region. Samples were finely ground and mixed with spectroscopic-grade potassium bromide (KBr) to form transparent pellets under high pressure [20,34]. Spectra were collected in transmission mode over the spectral range of 4000 to 400 cm−1, with a resolution of 4 cm−1 and an average of 32 scans per sample to improve the signal-to-noise ratio. Characteristic absorption bands corresponding to N–H, C=O, C=N, and M–N (metal–nitrogen) or M–O (metal–oxygen) vibrations were analyzed to verify the successful formation of coordination bonds and the structural integrity of the frameworks.

2.2.2. Thermal Stability

Thermal stability of the synthesized coordination compounds was evaluated using thermogravimetric analysis (TGA) to determine their decomposition profile, thermal resistance, and structural robustness under increasing temperature. The analysis was conducted using a TG-SDTA 851e analyzer (Mettler Toledo, Carrer del Segrià, 7-9, 08940 Cornellà de Llobregat, Barcelona, Spain), which integrates thermogravimetry (TG) and simultaneous differential thermal analysis (SDTA), allowing for the detection of both mass loss and thermal events such as phase transitions or decomposition reactions. The measurements were carried out in dynamic mode under a nitrogen atmosphere to prevent unwanted oxidation during heating. Approximately 5–10 mg of each powdered sample was placed in an alumina crucible and subjected to a controlled heating program from room temperature (25 °C) to 1000 °C at a constant heating rate of 5 °C/min [35]. The resulting thermograms were analyzed to identify characteristic weight loss steps associated with processes such as desorption of surface-adsorbed water, decomposition of organic ligands, and degradation of the coordination framework. These data were used to assess the thermal stability and structural integrity of the coordination compounds, which are critical properties for their potential application in electrochemical systems operating under variable thermal conditions [34,36].

2.2.3. X-Ray Diffraction (XRD)

X-ray diffraction (XRD) analysis was carried out to investigate the crystallographic structure, phase purity, and degree of crystallinity of the synthesized copper-based coordination compounds [20,37]. The XRD patterns of the CuHCF powders were recorded using a Bruker D8 Advance diffractometer (Billerica, MA, USA), equipped with a Cu Kα1 radiation source (λ = 1.5406 Å). The instrument was configured with a vertical Bragg–Brentano θ–2θ goniometer, operated at 40 kV and 30 mA, and coupled to a scintillation detector for enhanced signal sensitivity. Data acquisition was performed over a 2θ angular range of 10° to 70°, with a step size of 0.020° (2θ) and a dwell time of 3.0 s per step, ensuring high-resolution pattern collection. The diffraction peaks were analyzed to identify crystalline phases and to verify the formation of the expected PBA framework. Peak positions and intensities were compared with reference patterns to assess crystalline and structural order [37,38]. For CCCu, the powder XRD patterns were indexed to a monoclinic unit cell, consistent with the single-crystal X-ray diffraction (SC-XRD) results. This confirmed that the bulk material retained the same crystal symmetry and structural parameters as the isolated single crystals [20]. Detailed structural information, including unit cell dimensions, bond lengths, bond angles, and atomic coordinates, was obtained from SC-XRD analysis. The resulting Crystallographic Information Files (CIFs) were deposited in the Cambridge Crystallographic Data Centre (CCDC).

2.2.4. Scanning Electron Microscopy (SEM/EDS)

The morphological characterization of the surfaces was carried out using a field emission and variable pressure scanning electron microscope (FE-SEM, model ZEISS Sigma 360 VP). Prior to analysis, the samples were coated with a conductive layer of gold using a sputter coater system (model Q150R ES Plus, Quorum Technologies, East Sussex, England, UK), operated under high-vacuum conditions to enhance surface conductivity and minimize charge accumulation.
Observations were performed in variable pressure mode, with accelerating voltages ranging from 5 to 15 kV, depending on the required resolution. Micrographs were obtained at various magnifications to examine surface morphology, including textures, micrometric structures, and characteristic roughness patterns. Additionally, the SEM system was coupled with an energy-dispersive X-ray spectroscopy (EDX) microanalysis system (model AZtecLiveLite), equipped with a 40 mm2 active area SDD UltimMax analytical detector (Oxford Instruments, Halifax Rd, High Wycombe, UK). This system enabled qualitative and semi-quantitative analysis of the elemental composition in selected regions of the examined surfaces.

2.3. Corrosion Studies

Corrosion experiments were conducted to evaluate the electrochemical behavior of compounds CCCu and CuHCF in chloride-rich environments, simulating seawater conditions. The primary focus was on understanding the contributions of three key partial electrochemical reactions: the HER, the ORR, and the coordination compound oxidation reaction (CCOR). Modified carbon paste electrodes were fabricated and used as working electrodes (WE), each tested in triplicate.
The working electrode was fabricated using graphite wax as the support, and the CCCu or CuHCF material was deposited over modify carbon paste under manual compression. The preparation of the graphite paste electrode does not interfere with the electrochemical results. The mixture was then tightly packed into a Teflon sheath (4 mm diameter, 10 mm length), designed with perforations for electrical contact, allowing integration into a rotating disc electrode (RDE) system [39]. Electrochemical measurements were performed using linear sweep voltammetry (LSV) to obtain current density versus potential curves, which reveal the redox behavior and corrosion tendencies of the materials. The measured current was normalized by the geometric area of the working electrode to calculate the current density, without the need to perform IR compensation during electrochemical measurements. Experiments were carried out using a BASI/RDE-2 rotating electrode setup coupled with an Epsilon potentiostat/galvanostat. The electrochemical cell consisted of the following: carbon paste electrode used as an inert conductive support, onto which CCCu or CuHCF was immobilized as a surface-modifying layer and directly exposed to the electrolyte, platinum wire as counter electrode, and Ag/AgCl (4 M KCl, saturated) as reference electrode (RE). All potentials were converted and reported versus the standard hydrogen electrode (SHE) for consistency and comparison [40]. The measurements followed a potential sweep in the anodic direction from −1200 to 0 mVSHE, as described in prior studies [11], enabling simultaneous observation of cathodic and anodic processes. To ensure surface stabilization and reproducibility, each working electrode was preconditioned by holding at −1200 mVSHE for 30 s before the start of the potential scan. This methodology enabled the deconvolution of the total current response into the contributions from HER, ORR, and CCOR, which are critical for understanding degradation mechanisms and electrochemical stability in saline environments [24]. The 0.5 M NaCl test solution was prepared by using analytical-grade reagents and distilled water and constantly bubbled oxygen during the experiment.

2.4. Kinetic Corrosion Analysis

Non-linear fitting of polarization data was conducted using a superposition model based on mixed potential theory, as detailed in our previous work [39,41]. This approach considers that the total current density can be represented as the sum of the partial anodic and cathodic reactions occurring on the working electrode during the corrosion process, as presented in Equation (1). Under this consideration, when the total current density ( i ) is null, the cathodic and anodic current densities have the same magnitude [42,43].
i = i O 2 + i H 2 + i C C
where i represent the total current density, i O 2 and i H 2 denote the partial reduction current densities for ORR and HER, respectively, and i C C denotes the partial oxidation current density for CCCu or CuHCF [41].
Although experimental measurement of i O 2 , i H 2 and i C C is not directly feasible, their values can be deduced by considering kinetic expressions for each one. In this framework, the partial reactions for HER and CCOR are considered to follow a charge-transfer kinetic mechanism, while the ORR follows a mixed charge-diffusion kinetic mechanism, according to Equations (2)–(4) [39]. These considerations are based on the identification of individual control mechanisms, charge transfer or mass diffusion, and their combined effect, mixed charge-diffusion, which are well characterized in a potentiodynamic polarization curve [44]. Specifically, a continuous exponential increase in the current density with applied potential indicates a charge-transfer mechanism, whereas the presence of a planar plateau that gradually converges to the mixed potential/corrosion potential indicates a mixed charge-diffusion mechanism [44].
i O 2 = i 0 , O 2 1 − i O 2 i l , O 2 p e − 2.303 η O 2 t O 2
i H 2 = i 0 , H 2 e − 2.303 η H 2 t H 2
i C C = i 0 , C C e 2.303 η C C t C C
where the η O 2 = E − E e q , O 2 , η H 2 = E − E e q , H 2 and η C C = E − E e q C C are the ORR, HER, and CCOR, overpotentials, respectively; i 0 , O 2 , i 0 , H 2 , and i 0 , C C are the exchange current densities for ORR, HER, and CCOR, respectively; i l , O 2 is the limiting current density for ORR; t O 2 , t H 2 , and t C C are the Tafel slopes for ORR and HER and CCOR, respectively; E e q , O 2 , E e q , H 2 , and E e q C C are the equilibrium potentials for ORR, HER, and CCOR, respectively.

2.5. Computational Details

All calculations were carried out with Gaussian 16 using a representative fragment of the crystal structure rather than periodic models, since the reactivity is localized in the metal coordination environment [45]. Geometries, vibrational frequencies, and energies were optimized at the ωB97X-D/def2-SVP level with the Solvation Model based on Density (SMD) implicit solvation model (water), chosen for its accurate treatment of dispersion and reliable performance in calculating reaction barriers of coordination complexes. Natural Bond Orbital (NBO) charges, Overlap Population Density of States (OPDOS), and Reduced Density Gradient (RDG) analyses were performed with the open-source program Multiwfn 3.8 to characterize the electronic evolution throughout the reaction pathway [46].

3. Results and Discussion

3.1. IR Analysis

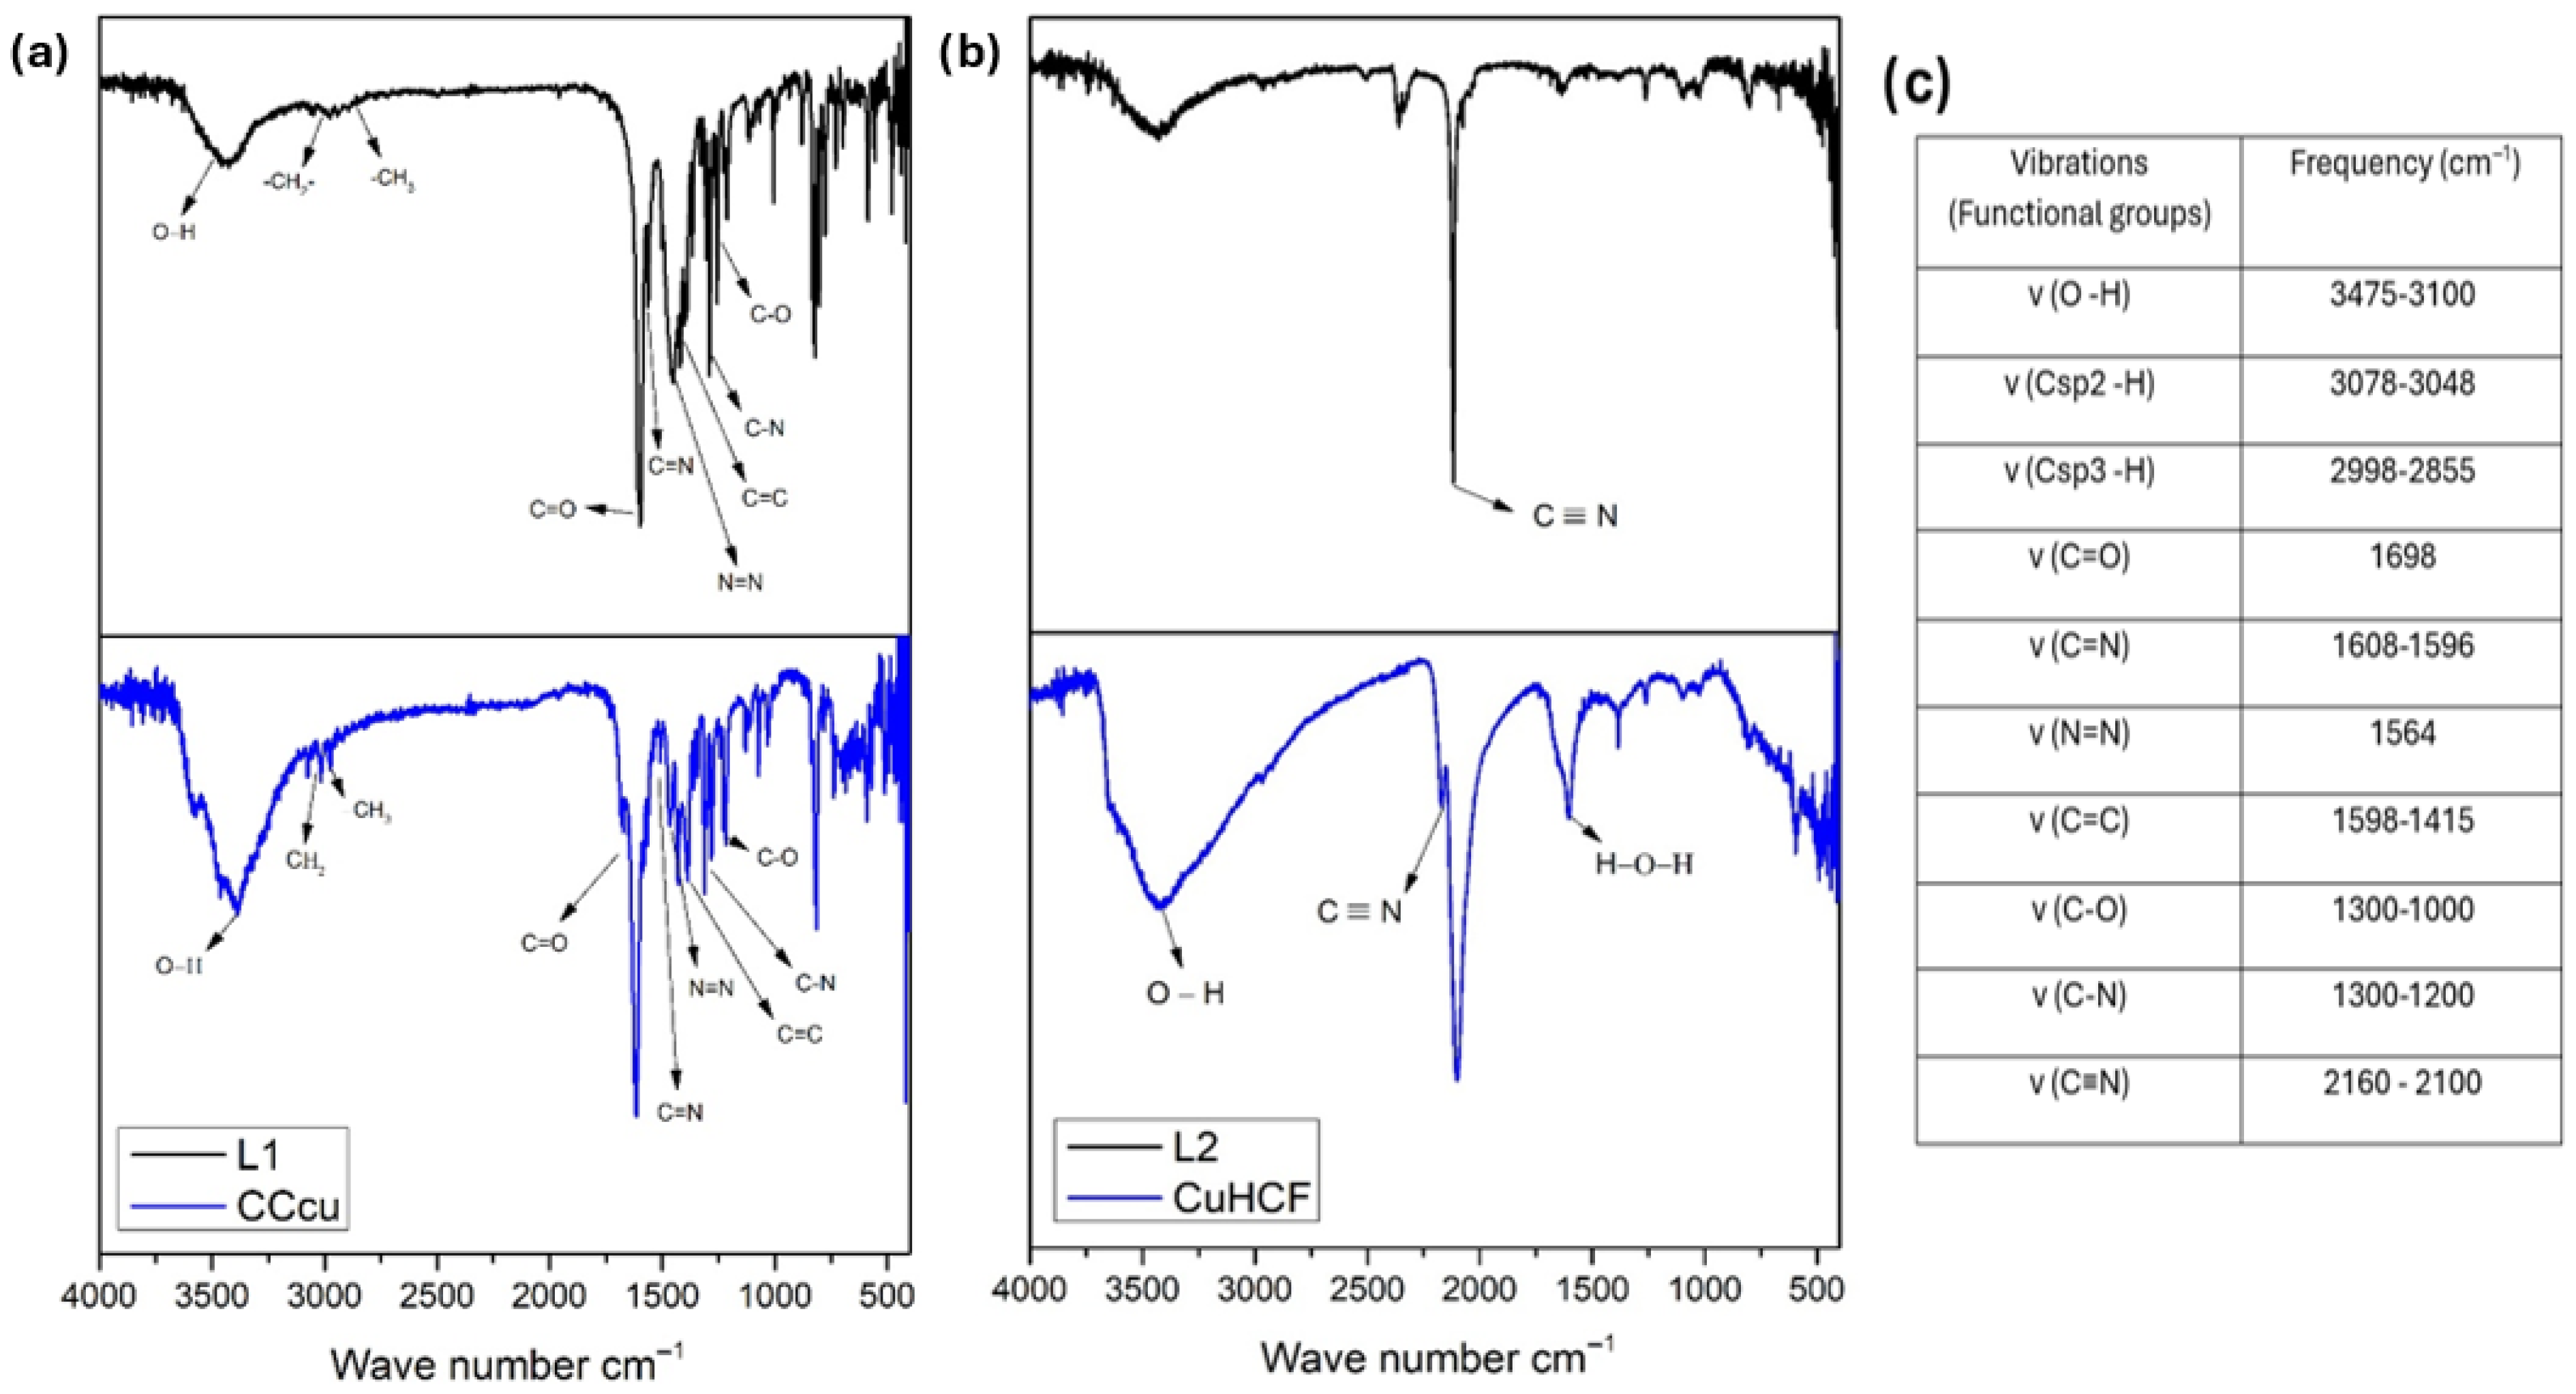
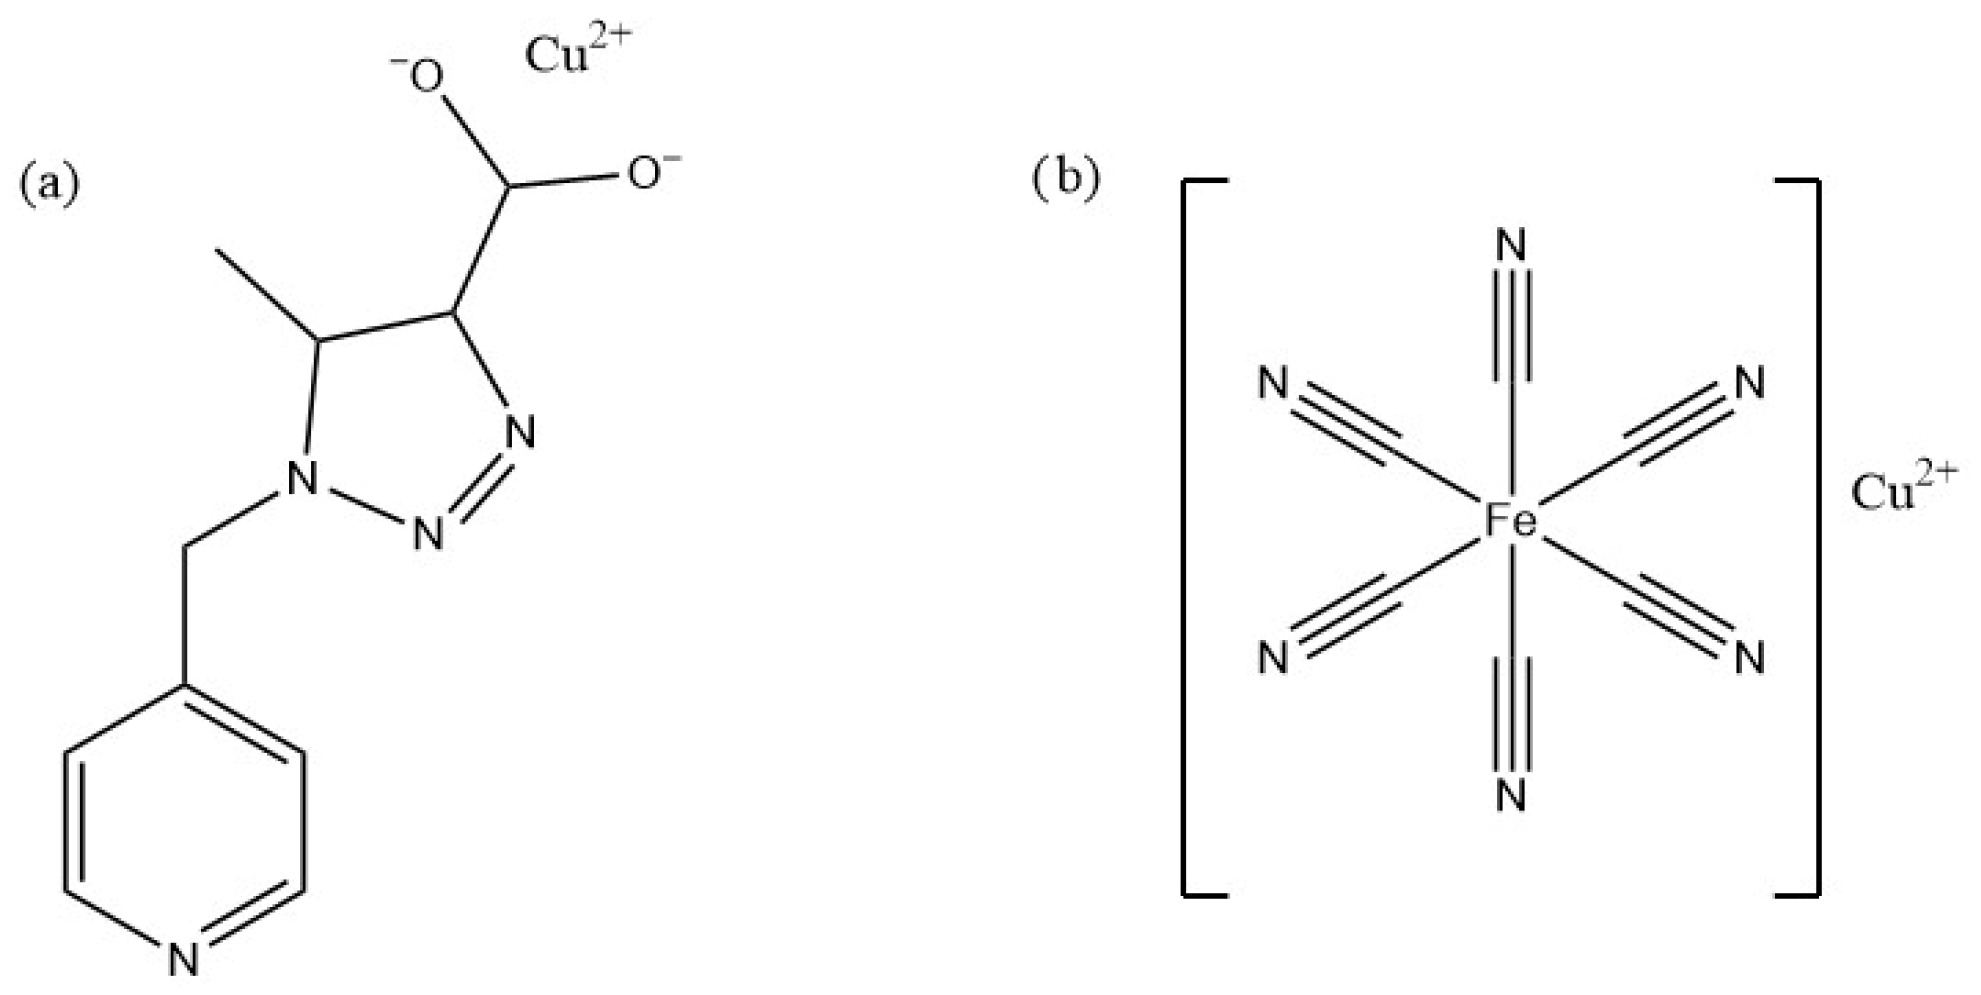
Figure 3 shows the IR spectroscopy, which was performed to characterize the functional groups present in the organic ligands and to confirm the formation of coordination bonds in the synthesized copper-based compounds. The analysis included both the organic ligand L1 and its corresponding coordination polymer (CCCu), as well as the ligand precursor [K3(Fe(CN)6)], referred to here as ligand L2, and its coordination product, CuHCF. The resulting spectra are presented in Figure 3a for L1 and CCCu, and Figure 3b for L2 and CuHCF [38,47]. The structures of the ligands and their coordination modes are illustrated in Figure 4, which serves as a guide for interpreting the assignments. Figure 3c summarizes the stretching and vibrations of the systems studied.
The spectrum of L1 exhibits a broad vibrational band at ~3470 cm−1 attributable to the ν(O–H) stretching of water molecules. The aliphatic C–H vibrations appear as (–CH2–) at ~2925 cm−1 and (–CH3) at ~2870 cm−1. An intense vibration at ~1610 cm−1 corresponds to the ν(C=O) double bond; the ν(C=N) stretching of the pyridine ring is observed at ~1560 cm−1. At ~1470 cm−1, a vibration associated with the 1,2,3-triazole ring with a contribution from ν(N=N) is detected. Additionally, ν(C=C) vibrations corresponding to aromatic ring are recorded at ~1410–1430 cm−1, ν(C–N) at ~1250–1290 cm−1, and ν(C–O) at ~1150–1200 cm−1. In CCCu, the ligand structure is preserved, but characteristic shifts appear that agree with the proposed coordination shown in Figure 3a (chelation of Cu2+ via the carbonyl oxygen and nitrogen donors): the ν(C=O) shifts to ~1630–1670 cm−1 and the ν(C=N) of the pyridine core to ~1510–1530 cm−1. The vibrational band of the 1,2,3-triazole ring corresponding to ν(N=N) is located at ~1430 cm−1 and shows a slight shift to lower wavenumber relative to L1, consistent with its involvement in the coordination environment. The ν(O–H) contribution appears broader and more intense at ~3440 cm−1 than in L1, suggesting a greater fraction of incorporated water (coordinated and/or crystallization) within the polymeric matrix. Ligand L2 (K3[Fe(CN)6]) presents a sharp ν(C≡N) at ~2116 cm−1, characteristic of cyanide coordinated to Fe3+. In CuHCF, the ν(C≡N) stretching is observed at ~2090–2100 cm−1, shifted to a lower wavenumber relative to the precursor, in agreement with the interaction of the cyanide group with Fe3+ and Cu2+ centers within the Prussian Blue-type structure represented in Figure 3b. The CuHCF spectrum also shows a broad ν(O–H) vibrational band at ~3435 cm−1 and the δ(H–O–H) at ~1610–1615 cm−1, attributable to hydration water (coordinated and/or crystallization). In the low-frequency region, ν(Fe–C) at ~600–620 cm−1 and ν(Cu–N) at ~500–520 cm−1 are noteworthy; however, distinguishing both contributions clearly exceeds the resolution of these spectra, so they are only mentioned as being consistent with the expected structure.
Figure 4 presents the 2D chemical structures of copper-based compounds. In Figure 4a, CCCu is shown, consisting of a triazole-derived ligand with carboxylate groups coordinating to Cu2+. The structure features aromatic and heteroaromatic rings connected through the triazole moiety, which provides chelation sites for copper. In Figure 4b, CuHCF is illustrated as a cubic coordination framework, where Fe is octahedrally surrounded by six cyanide (–C≡N) ligands, forming a [Fe(CN)6]4− unit electrostatically balanced by Cu2+ cations. Together, these representations highlight the structural diversity of coordination chemistry applied to copper complexes, with CCCu serving as an organic–inorganic hybrid and CuHCF as a Prussian Blue analogue featuring an extended inorganic framework.
The vibrational bands ν(O–H) at ~3435 cm−1 and δ(H–O–H) at ~1612 cm−1 confirm the presence of hydrated domains within the CuHCF solid, while the shift of ν(C≡N) to ~2090 cm−1 reflects the dual Fe/Cu metallic environment of the cyanide. Taken together, these features indicate a porous, hydrated inorganic coordination network with more direct ionic access and electronic pathways through the cyanide ligand than in CCCu. In contrast, in CCCu, the chelating coordination, evidenced by the shifts in ν(C=O) and ν(C=N) along with the persistence of the 1,2,3-triazole ring band, is consistent with a more saturated and rigid organic coordination framework around Cu2+, which reduces solvent accessibility to the metal center. In this context, the FTIR evidence suggests that CuHCF may exhibit faster kinetics in processes associated with hydrogen evolution under cathodic polarization compared to CCCu.
The FTIR spectroscopic analysis confirms the successful coordination of the organic ligands to copper centers in both CCCu and CuHCF. The observed shifts in key vibrational bands, particularly the C=O and N=N modes in CCCu and the C≡N band in CuHCF, demonstrate the formation of stable metal–ligand interactions, thereby validating the structural integrity of the coordination networks. The presence of coordinated or crystallization water, evidenced by broad O–H stretching bands, further supports the incorporation of hydration into the polymeric and porous frameworks. These findings establish the chemical identity and coordination environment of the synthesized compounds, providing a robust foundation for subsequent electrochemical and corrosion resistance evaluations.

3.2. Thermogravimetric Analysis

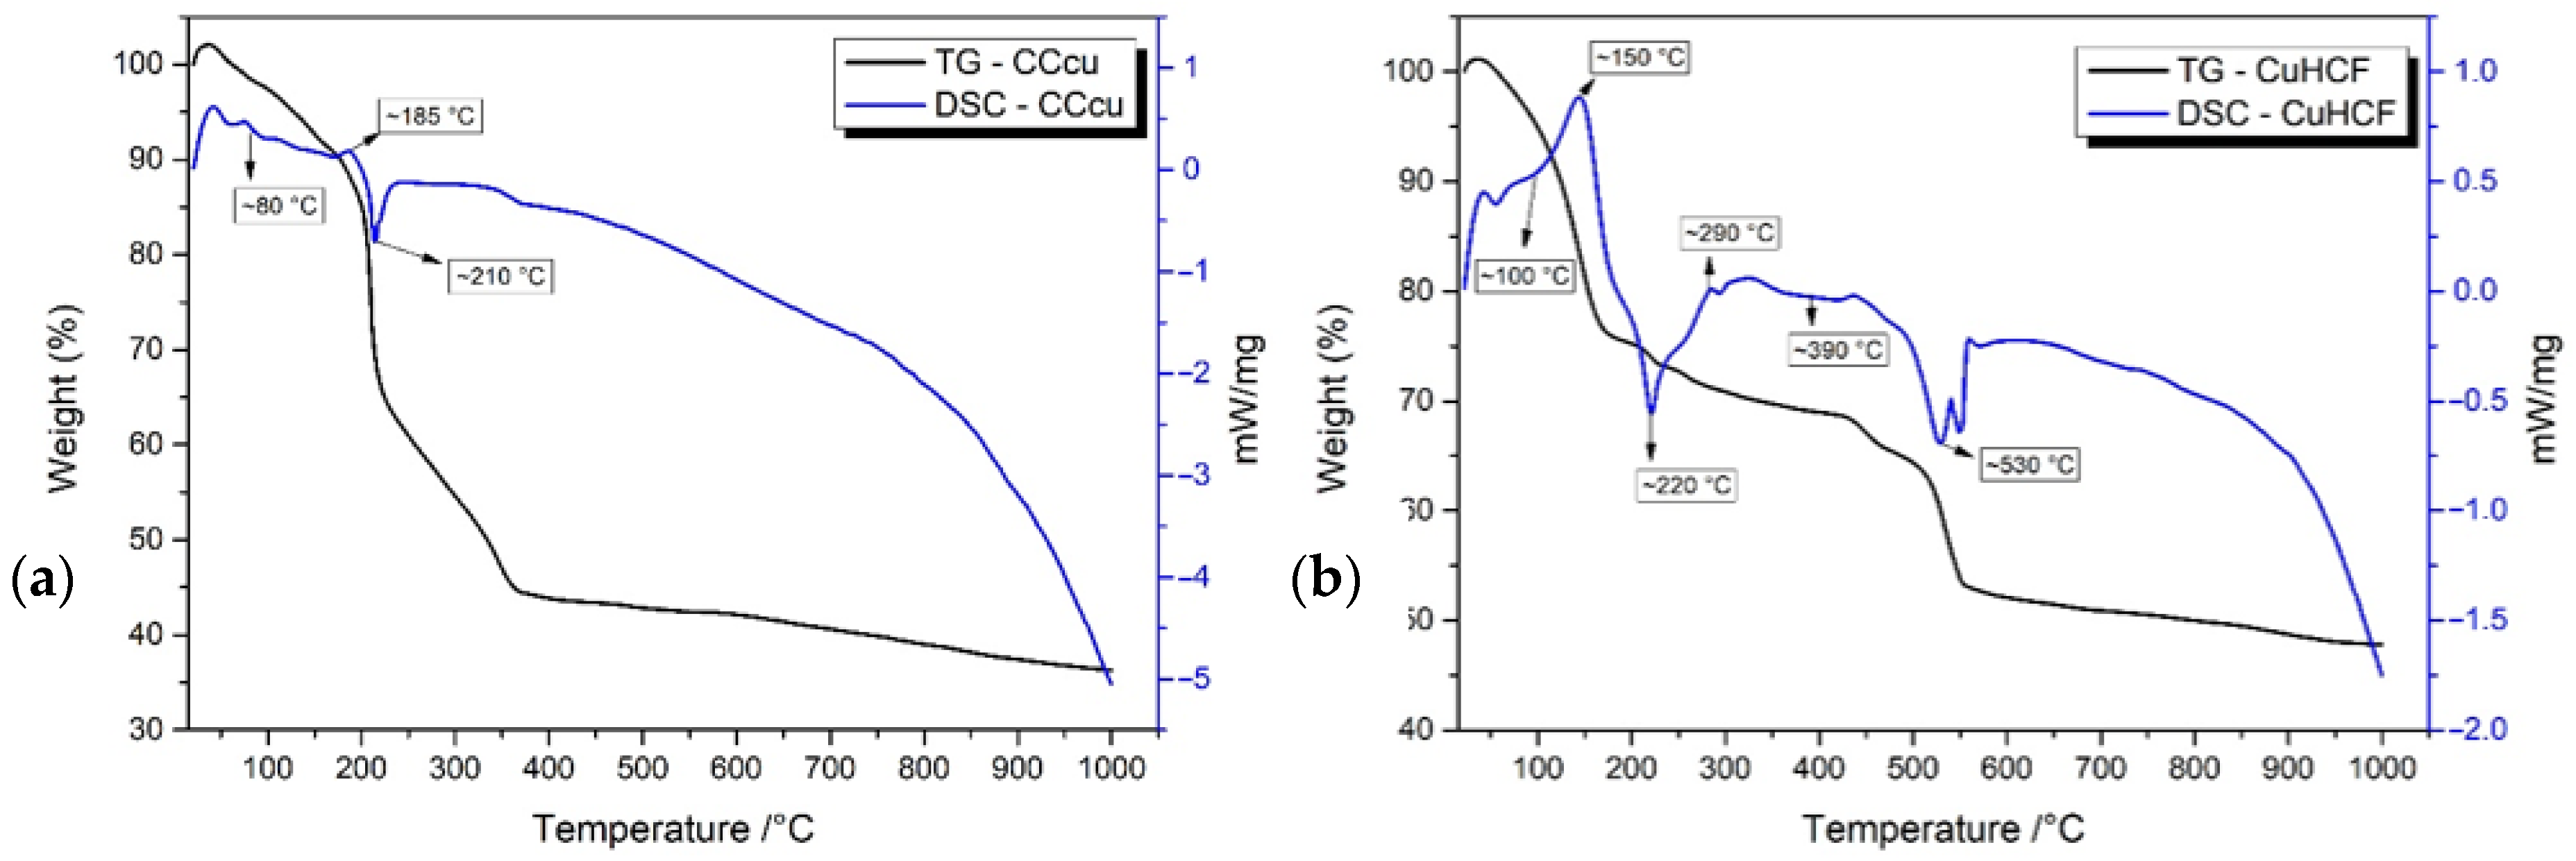
TGA coupled with DSC was conducted to evaluate the thermal stability and decomposition behavior of the synthesized coordination compounds. The measurements were carried out over the temperature range of 25 °C to 1000 °C, and the resulting curves are presented in Figure 5, where Figure 5a corresponds to CCCu and Figure 5b to CuHCF. Beyond describing mass and heat-flow events, TGA/DSC informs how each solid will behave in humid air and during operation at near-room temperature. In particular, the low-temperature endotherms and the first mass-loss steps reveal the presence of two water populations: (i) surface or weakly bound water, which desorbs below ~120 °C and governs short-term humidity uptake/release; and (ii) structural water, which participates in hydrogen-bonded bridges that stabilize and “stitch” neighboring nodes, enabling 2D expansion of the polymeric network and providing continuous proton/ion transport pathways. Mapping the onset, peak, and end set of these events defines the thermal safety window in which the framework can retain the water that supports its electrochemical function without undergoing dehydration-induced collapse.
In Figure 5a, the TGA curve of CCCu shows an initial mass loss of approximately 5% between 25 °C and 120 °C, accompanied by a mild endothermic peak in the DSC curve centered at ~80 °C. This process could be attributed to the desorption of physically adsorbed and/or weakly coordinated moisture, whose release is endothermic due to the breaking of hydrogen bonds with the matrix, a behavior commonly observed in coordination networks with N/O donor atoms, as reported in the recent literature on CPs and MOFs [48]. For CCCu, the weak endotherm at ~80 °C is consistent with surface/weakly bound water only, while no distinct higher-temperature endotherm attributable to persistent structural water is resolved. This suggests that electrochemical stability at ambient conditions will rely mainly on physisorbed moisture, with limited network reinforcement by water bridges; accordingly, modest dehydration, even at mild heating can lead to faster loss of ligand integrity and a narrower operating window before structural decay initiates. Above 120 °C, the TGA profile enters a region of significant mass loss (~50%) up to ~350 °C, likely associated with the decomposition of the organic ligands (pyridine, triazole, and carboxylate). Correspondingly, the DSC shows a well-defined exothermic peak between 200 and 220 °C (Tₚ ≈ 210 °C), consistent with combustion and oxidative fragmentation of the organic ligand framework, a behavior previously reported for CPs of this type [20,49]. In the 120–185 °C interval, the DSC reveals a mixed thermal event, starting slightly with an endothermic process and then shifting to exothermic, suggesting partial structural reorganization or ligand destabilization prior to the main decomposition. From 350 °C to 1000 °C, the TGA curve exhibits an additional gradual mass loss (~5%), while the DSC presents a broad exothermic tail attributable to the slow oxidation of residual carbonaceous material and the progressive formation of copper oxides. Overall, CCCu exhibits limited thermal stability in the low-to-medium temperature range (≤350 °C). Therefore, for applications requiring structural integrity, operating temperatures should be maintained below 120 °C. In Figure 5b, the TGA curve of CuHCF shows a higher initial mass loss (~8%) between 25 °C and 150 °C, accompanied by a significant endothermic peak in the DSC curve starting around 100 °C and reaching its maximum at ~150 °C. This is attributable to the desorption of physically adsorbed and/or weakly coordinated moisture. In contrast, the broader initial endotherm of CuHCF (100–150 °C), together with the subsequent two-step pattern (exo at ~220 °C and endo at ~290 °C), supports the presence of structural water that acts as a supramolecular binder between CN-linked nodes. Such H-bonded water bridges promote planar (2D) expansion of the Prussian-Blue-type network and help maintain percolating ionic pathways, which is beneficial for charge transfer and interfacial wetting under ambient electrochemical operation. TGA/DSC thus provides an indirect yet robust confirmation of this structural-water contribution and delineates temperature thresholds beyond which dehydration triggers framework softening or collapse, impairing electrochemical performance. In PBAs, initial desorption stages of weakly bound species are reported prior to the decomposition of CN bridges [50]. Between 150 and 350 °C, a moderate mass loss (~15%) is observed; in this interval, the DSC displays a pronounced exothermic signal beginning at ~160 °C with a well-defined maximum at ~220 °C, followed by an endothermic peak around ~290 °C. This pattern suggests a two-step process: (i) collapse/reorganization of the Cu–C≡N–Fe framework with heat release, and (ii) an endothermic step associated with deeper cleavage of cyanide (C≡N) bonds within the lattice. This staged energy demand in PB/PBA degradation under oxidative atmospheres has been documented with reactive windows around ~200–220 °C, marking the onset of oxidation [51].
From 350 °C to 600 °C, CuHCF undergoes its most pronounced thermal degradation (~30%), and the DSC curve exhibits multiple exothermic peaks, consistent with the oxidation of C/N residues and the formation of metal oxides (Fe2O3, CuO, or mixed oxides) [52]. Above 600 °C, the mass stabilizes, and the DSC profile shows a weaker exothermic tail, indicating the completion of decomposition and the formation of a thermally stable residue composed of mixed oxides. Taken together, the TG/DSC evidence indicates that CCCu exhibits limited low-to-medium-temperature stability with negligible structural-water support, so maintaining T < ~120 °C is essential to preserve ligand integrity and avoid rapid capacity fade. CuHCF, in turn, shows broader water-mediated stabilization and more gradual dehydration, which translates into better retention of the conductive/ionic scaffold at room temperature and moderate heating. Defining these dehydration/decomposition limits is therefore directly relevant for predicting performance, durability, and safe operating ranges in ambient electrochemical applications. CuHCF demonstrates greater thermal stability than CCCu: its metal–cyanide framework remains essentially intact up to ~150 °C and begins to degrade more significantly above 250 °C, with complete decomposition occurring above 600 °C. Combined with its greater water retention/release control, which favors ionic transport, CuHCF emerges as a more suitable material for prolonged operation at elevated temperatures and for systems where rapid electrochemical activation is desirable; by contrast, CCCu is more appropriate for environments with moderate thermal exposure.

3.3. X-Ray Diffraction (XRD) Analysis

3.3.1. CCCu Material

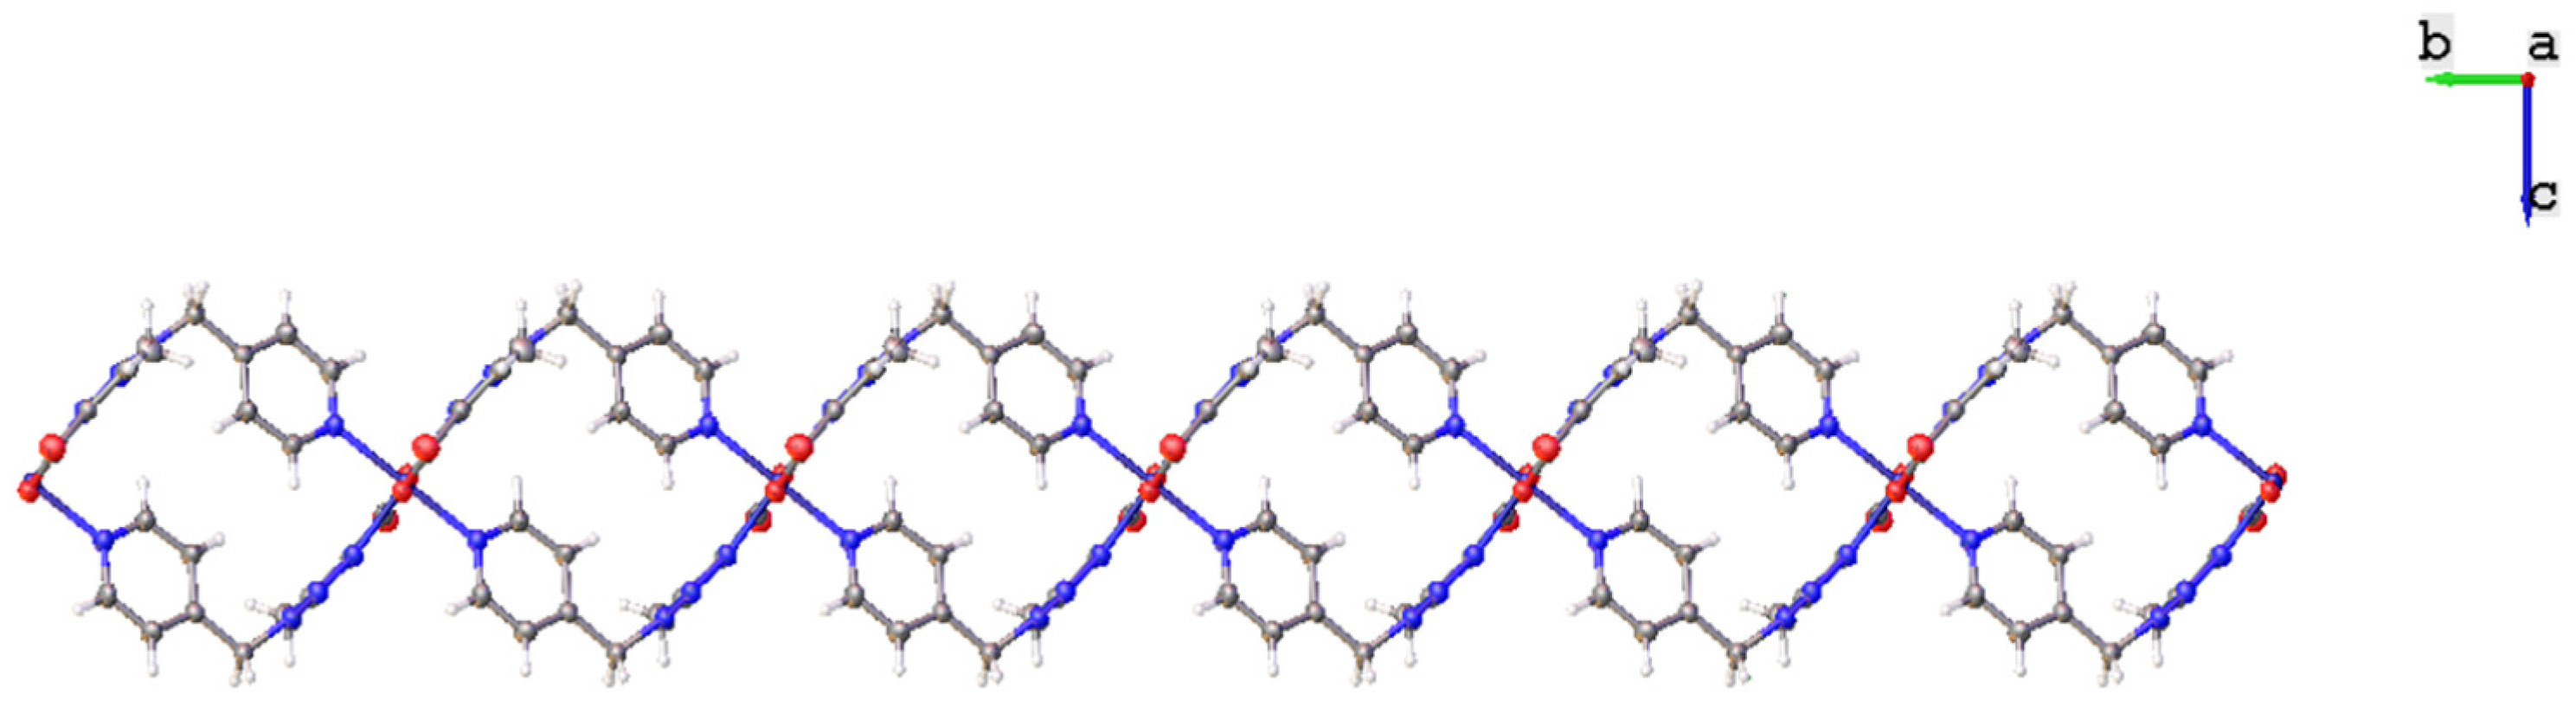
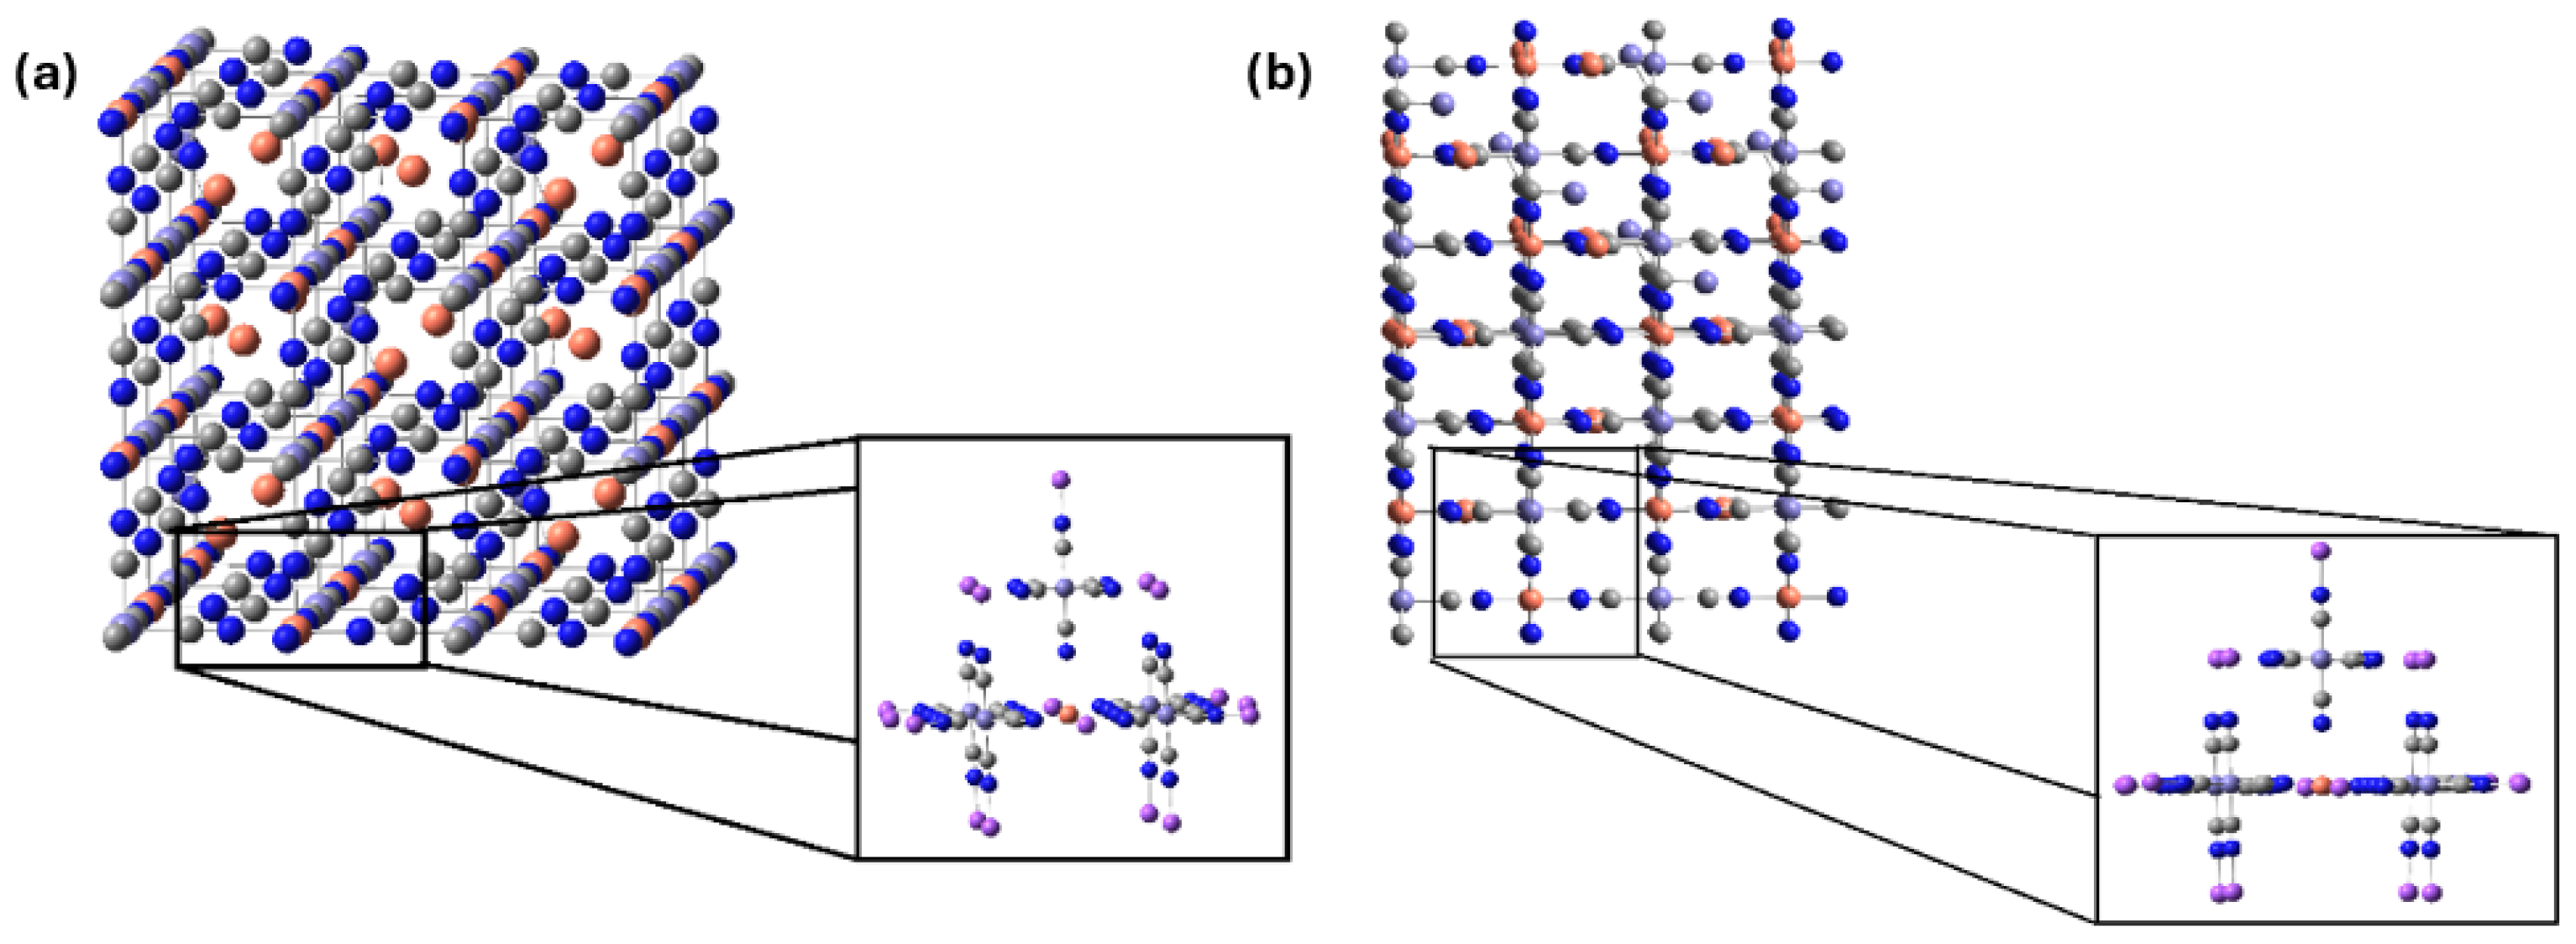
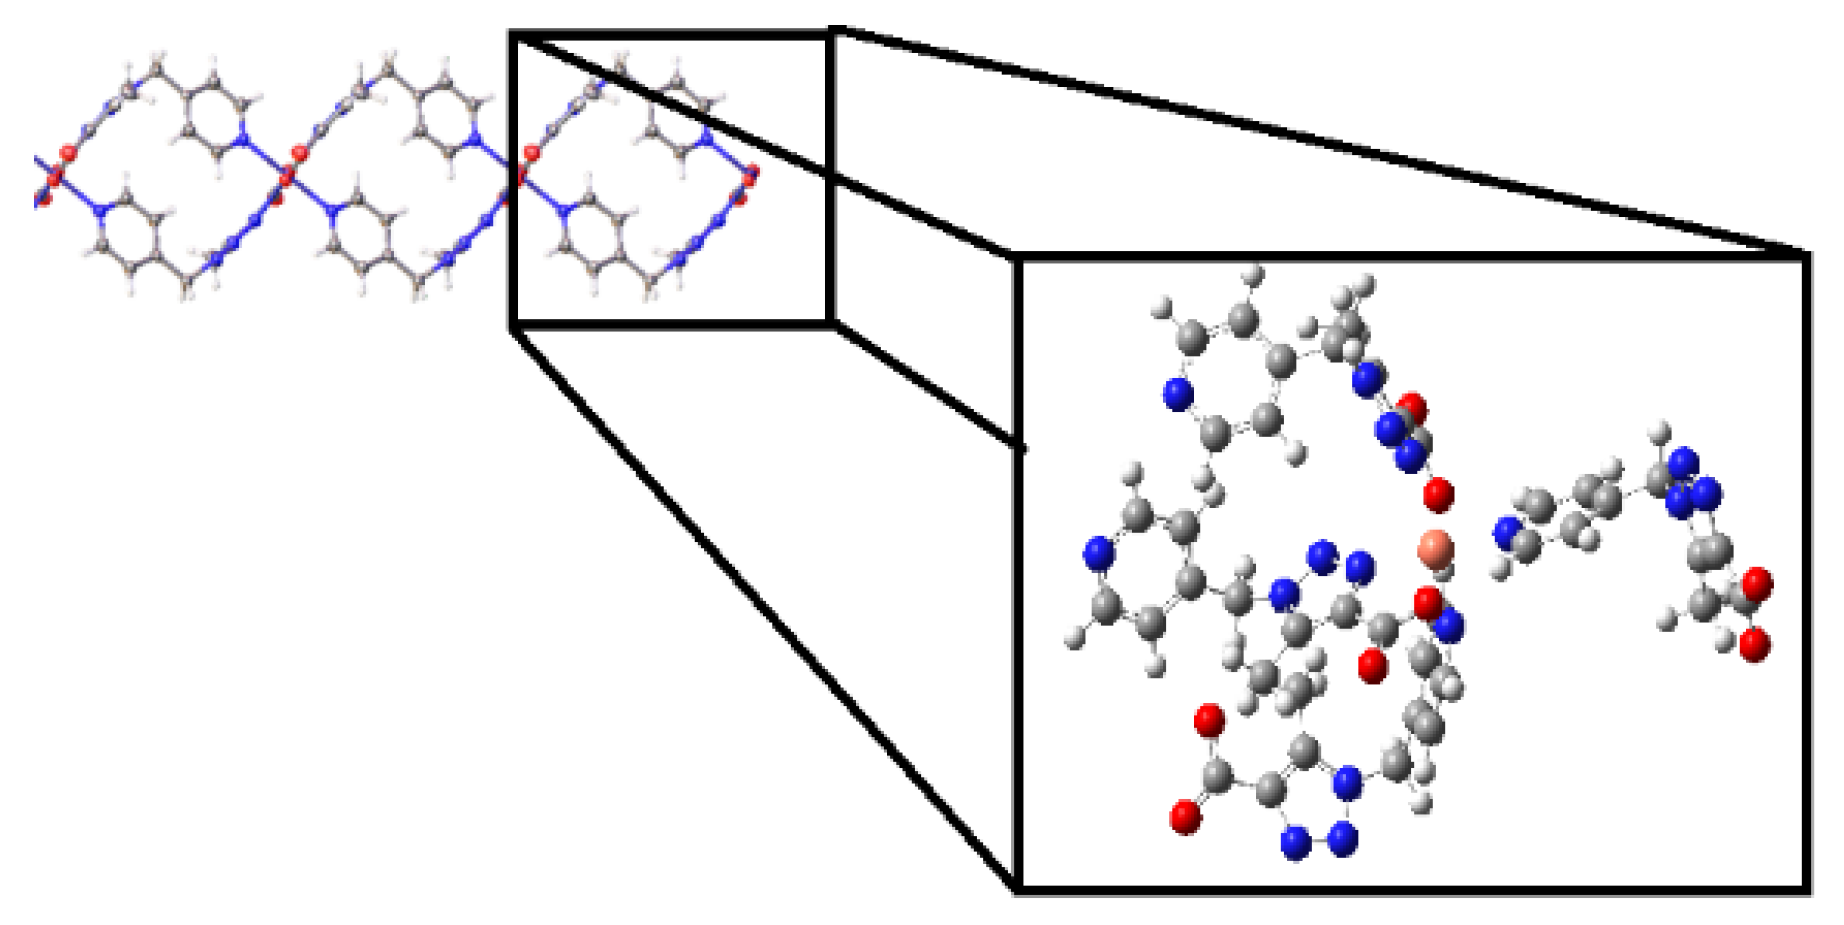
The structure of the coordination compound CCCu is shown in Figure 6 and Figure 7. The distorted octahedral geometry of CCCu, defined by a cis-N4O2 coordination environment (with Cu–N and Cu–O bond lengths of approximately 2.03 Å and 2.40 Å, respectively), creates a favourable electronic structure that promotes efficient overlap between the metal d-orbitals and the π-orbitals of the nitrogen-containing ligands. This electronic configuration is particularly advantageous for enhancing electron transfer processes relevant to HER. As reported by Delgado et al. [30], in coordination polymers based on triazol–pyridinium ligands, such π–metal interactions significantly improve intrinsic conductivity and catalytic performance by lowering the activation energy required for H–H bond formation.
When compared with the well-studied CuHCF material used as an HER electrocatalyst and H2O2 sensor, CCCu exhibits analogous electrochemical behavior with distinct distinctions. Wessells et al. [38] report that Prussian Blue analogs such as CuHCF achieve high currents and low overpotentials (~100 mVSHE at 10 mA·cm−2) thanks to their open framework and Fe vacancies that facilitate ionic diffusion. However, CCCu features a 2D supramolecular hydrogen-bond network that creates discrete proton channels while protecting Cu2+ from dissolution via its organic ligands. This suggests a potential balance between ionic conductivity and metal protection superior to CuHCF, where the open structure exposes more active sites but also is more vulnerable to Cu leaching during charge–discharge cycling.
Moreover, electrochemical studies on CuHCF show that although the theoretical capacity of cyanoferrate analogs is high, their long-term stability is compromised by the expansion and contraction of the PBA lattice during ion insertion [53]. In contrast, CCCu is rigid, well-defined Cu–N and Cu–O coordination environment minimizes volumetric changes, resulting in reduced structural degradation under alternating potentials.
In terms of corrosion resistance, CCCu exhibits intrinsic protection: the triazole and pyridyl ligands strongly encapsulate the metal, minimizing direct exposure to the electrolyte. CCCu, on the other hand, achieves synthetic simplicity: a single coordination step crystallizes both the protective and conductive network simultaneously. CCCu not only inherits the high catalytic activity of Cu2+ coordinated in π-conjugated environments but also achieves superior structural and chemical resilience compared to many of its analogs, offering greater stability and reduced Cu2+ leaching versus CuHCF. This balance of mixed conductivity, corrosion protection, and ease of manufacture positions CCCu as a promising candidate for green-hydrogen electrodes in solar-driven electrolysis and justifies future kinetic tests and prolonged cycling studies to validate its performance under real operating conditions.

3.3.2. CuHCF Material

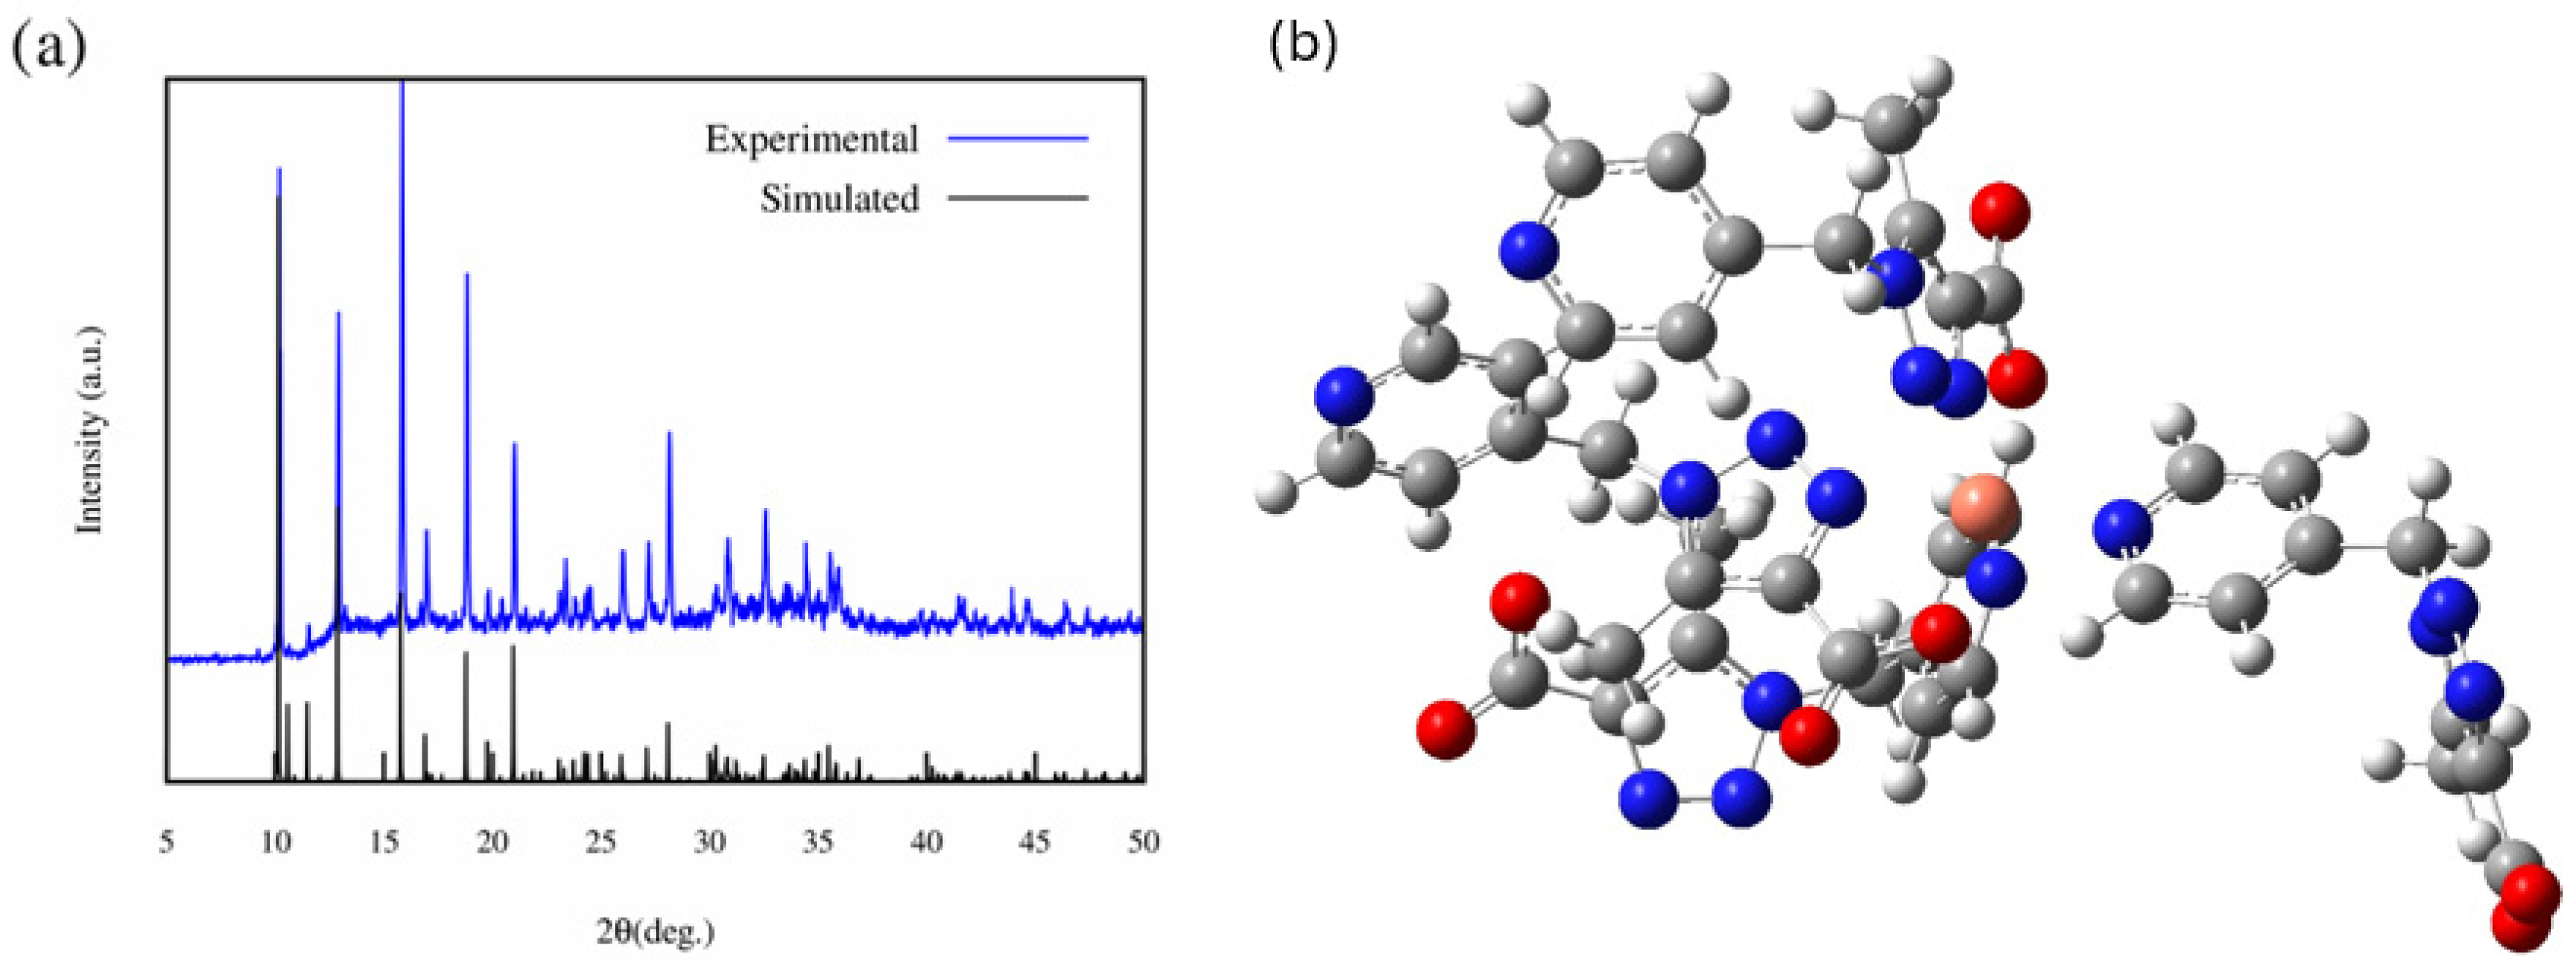
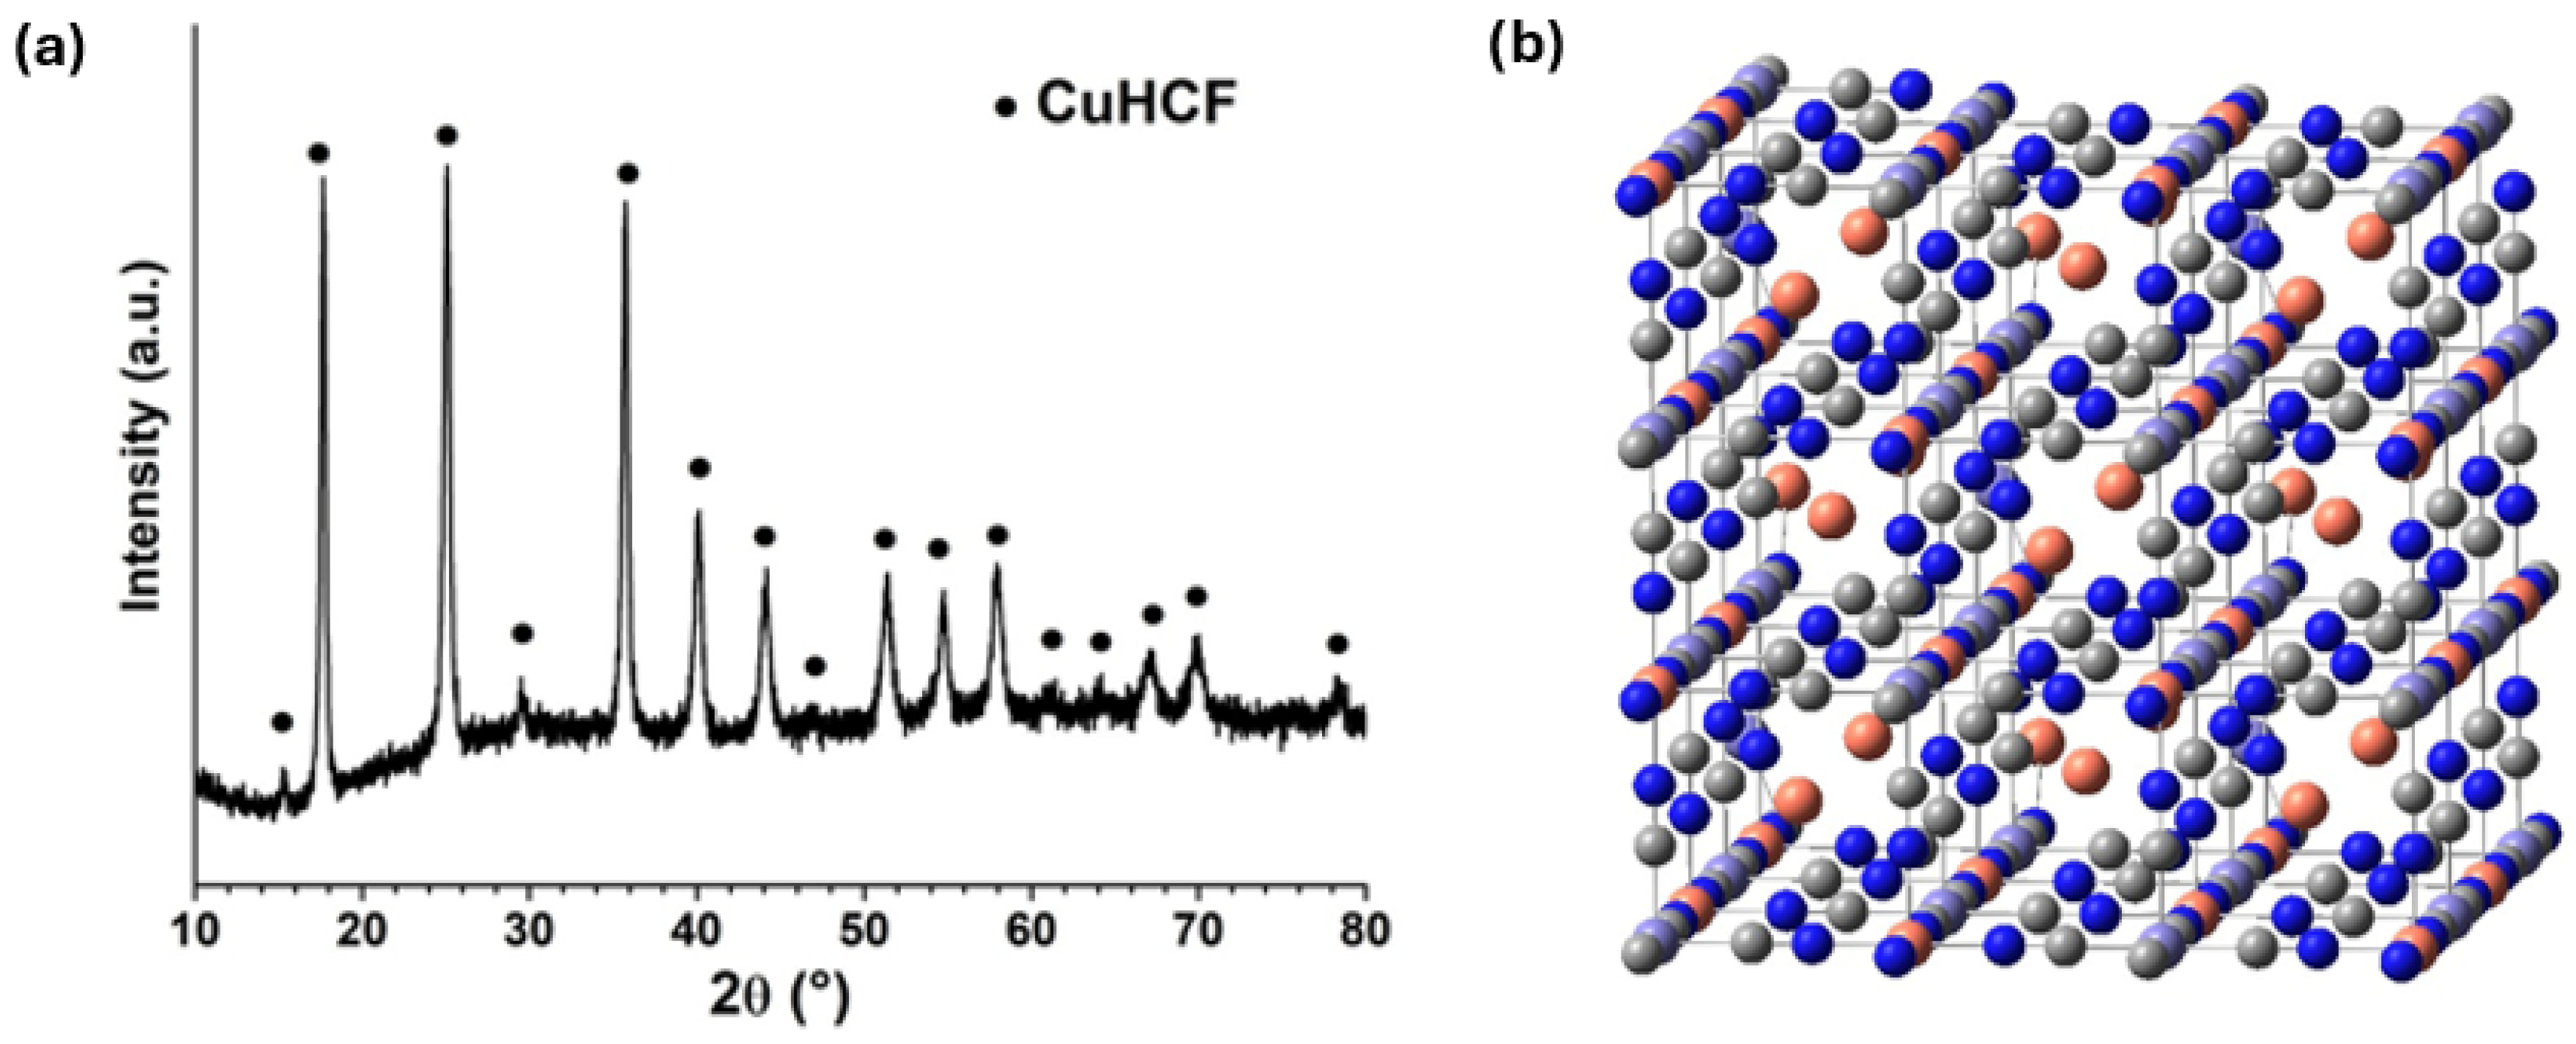
Figure 8 shows the structural characterisation of copper hexacyanoferrate. Figure 8a presents the powder XRD pattern, where the intense and sharp peaks confirm the high crystallinity of the material and match the reflections of a cubic PBA structure. Figure 8b illustrates the crystallographic model of CuHCF, highlighting the three-dimensional open framework formed by Cu and Fe centres bridged by cyanide ligands (–C≡N–), which creates porous channels capable of hosting cations and water molecules. Together, these results demonstrate the successful synthesis of CuHCF with an ordered crystalline structure and the porous network typical of PBAs, underpinning its potential for applications in energy storage and electrocatalysis.
Powder XRD characterization of the copper-based PBA Figure 8a, specifically Cu1.333[Fe(CN)6]0.667·H2O (hereafter CuHCF), reveals a face-centered cubic lattice (space group Fm-3m) with a lattice parameter a = 10.07 Å, calculated from the principal peaks at 2θ = 17.68°, 25.10°, 35.74°, 39.78°, and 43.66°. These reflections correspond to the (200), (220), (400), (420), and (422) planes, respectively, confirming the crystalline and phase purity of the synthesized material [54]. Each Cu2+ ion is octahedrally coordinated to six nitrogen atoms of Fe(CN)62−, yielding a robust three-dimensional framework whose approximately 33% vacancies are charge-balanced by water molecules, forming hydrogen-bond bridges and interstitial channels that facilitate proton conduction. The crystal structure of the coordination compound CuHCF is depicted in Figure 8b.
The highly ordered framework of CuHCF, composed of a three-dimensional network of Cu2+ ions octahedrally coordinated to Fe(CN)62− units, establishes a uniform array of equidistant redox-active sites. This crystallographic regularity facilitates rapid and isotropic electron transport through equivalent pathways, thereby minimizing internal resistive losses. Recent studies have demonstrated that Prussian Blue analogues with high structural crystallinity and lattice parameters around 10 Å exhibit pronounced pseudocapacitive behaviour and narrow peak separations (ΔEₚ < 40 mV) in cyclic voltammetry, a consequence of the homogeneous distribution and accessibility of their redox centres [55]. The Fe(CN)62− vacancies, evidenced by slight asymmetry and relative intensity differences between the (220) and (420) peaks, indicate interstitial channels that accommodate water molecules. These interconnected 3D tunnels provide ideal pathways for H+ or Na+ migration from seawater. Rapid ion diffusion in PBA materials with similar unit cells has been confirmed, showing diffusion resistance below 5 Ω cm2 [33]. An XRD pattern free of secondary peaks ensures the absence of CuO/Cu2O phases, which could be responsible for rapid degradation in chloride-rich environments. The phase purity and the high density of Fe–C≡N–Cu linkages reinforced by cubic symmetry impart a physical and chemical barrier against Cu2+ leaching. Correspondingly, PBA materials with equivalent crystallinity exhibit corrosion rates in 0.5 M NaCl that are ten times lower than those of pure metallic copper, as demonstrated by linear polarization and weight-loss experiments [37].

3.4. Morphological Surface and Elemental Mapping

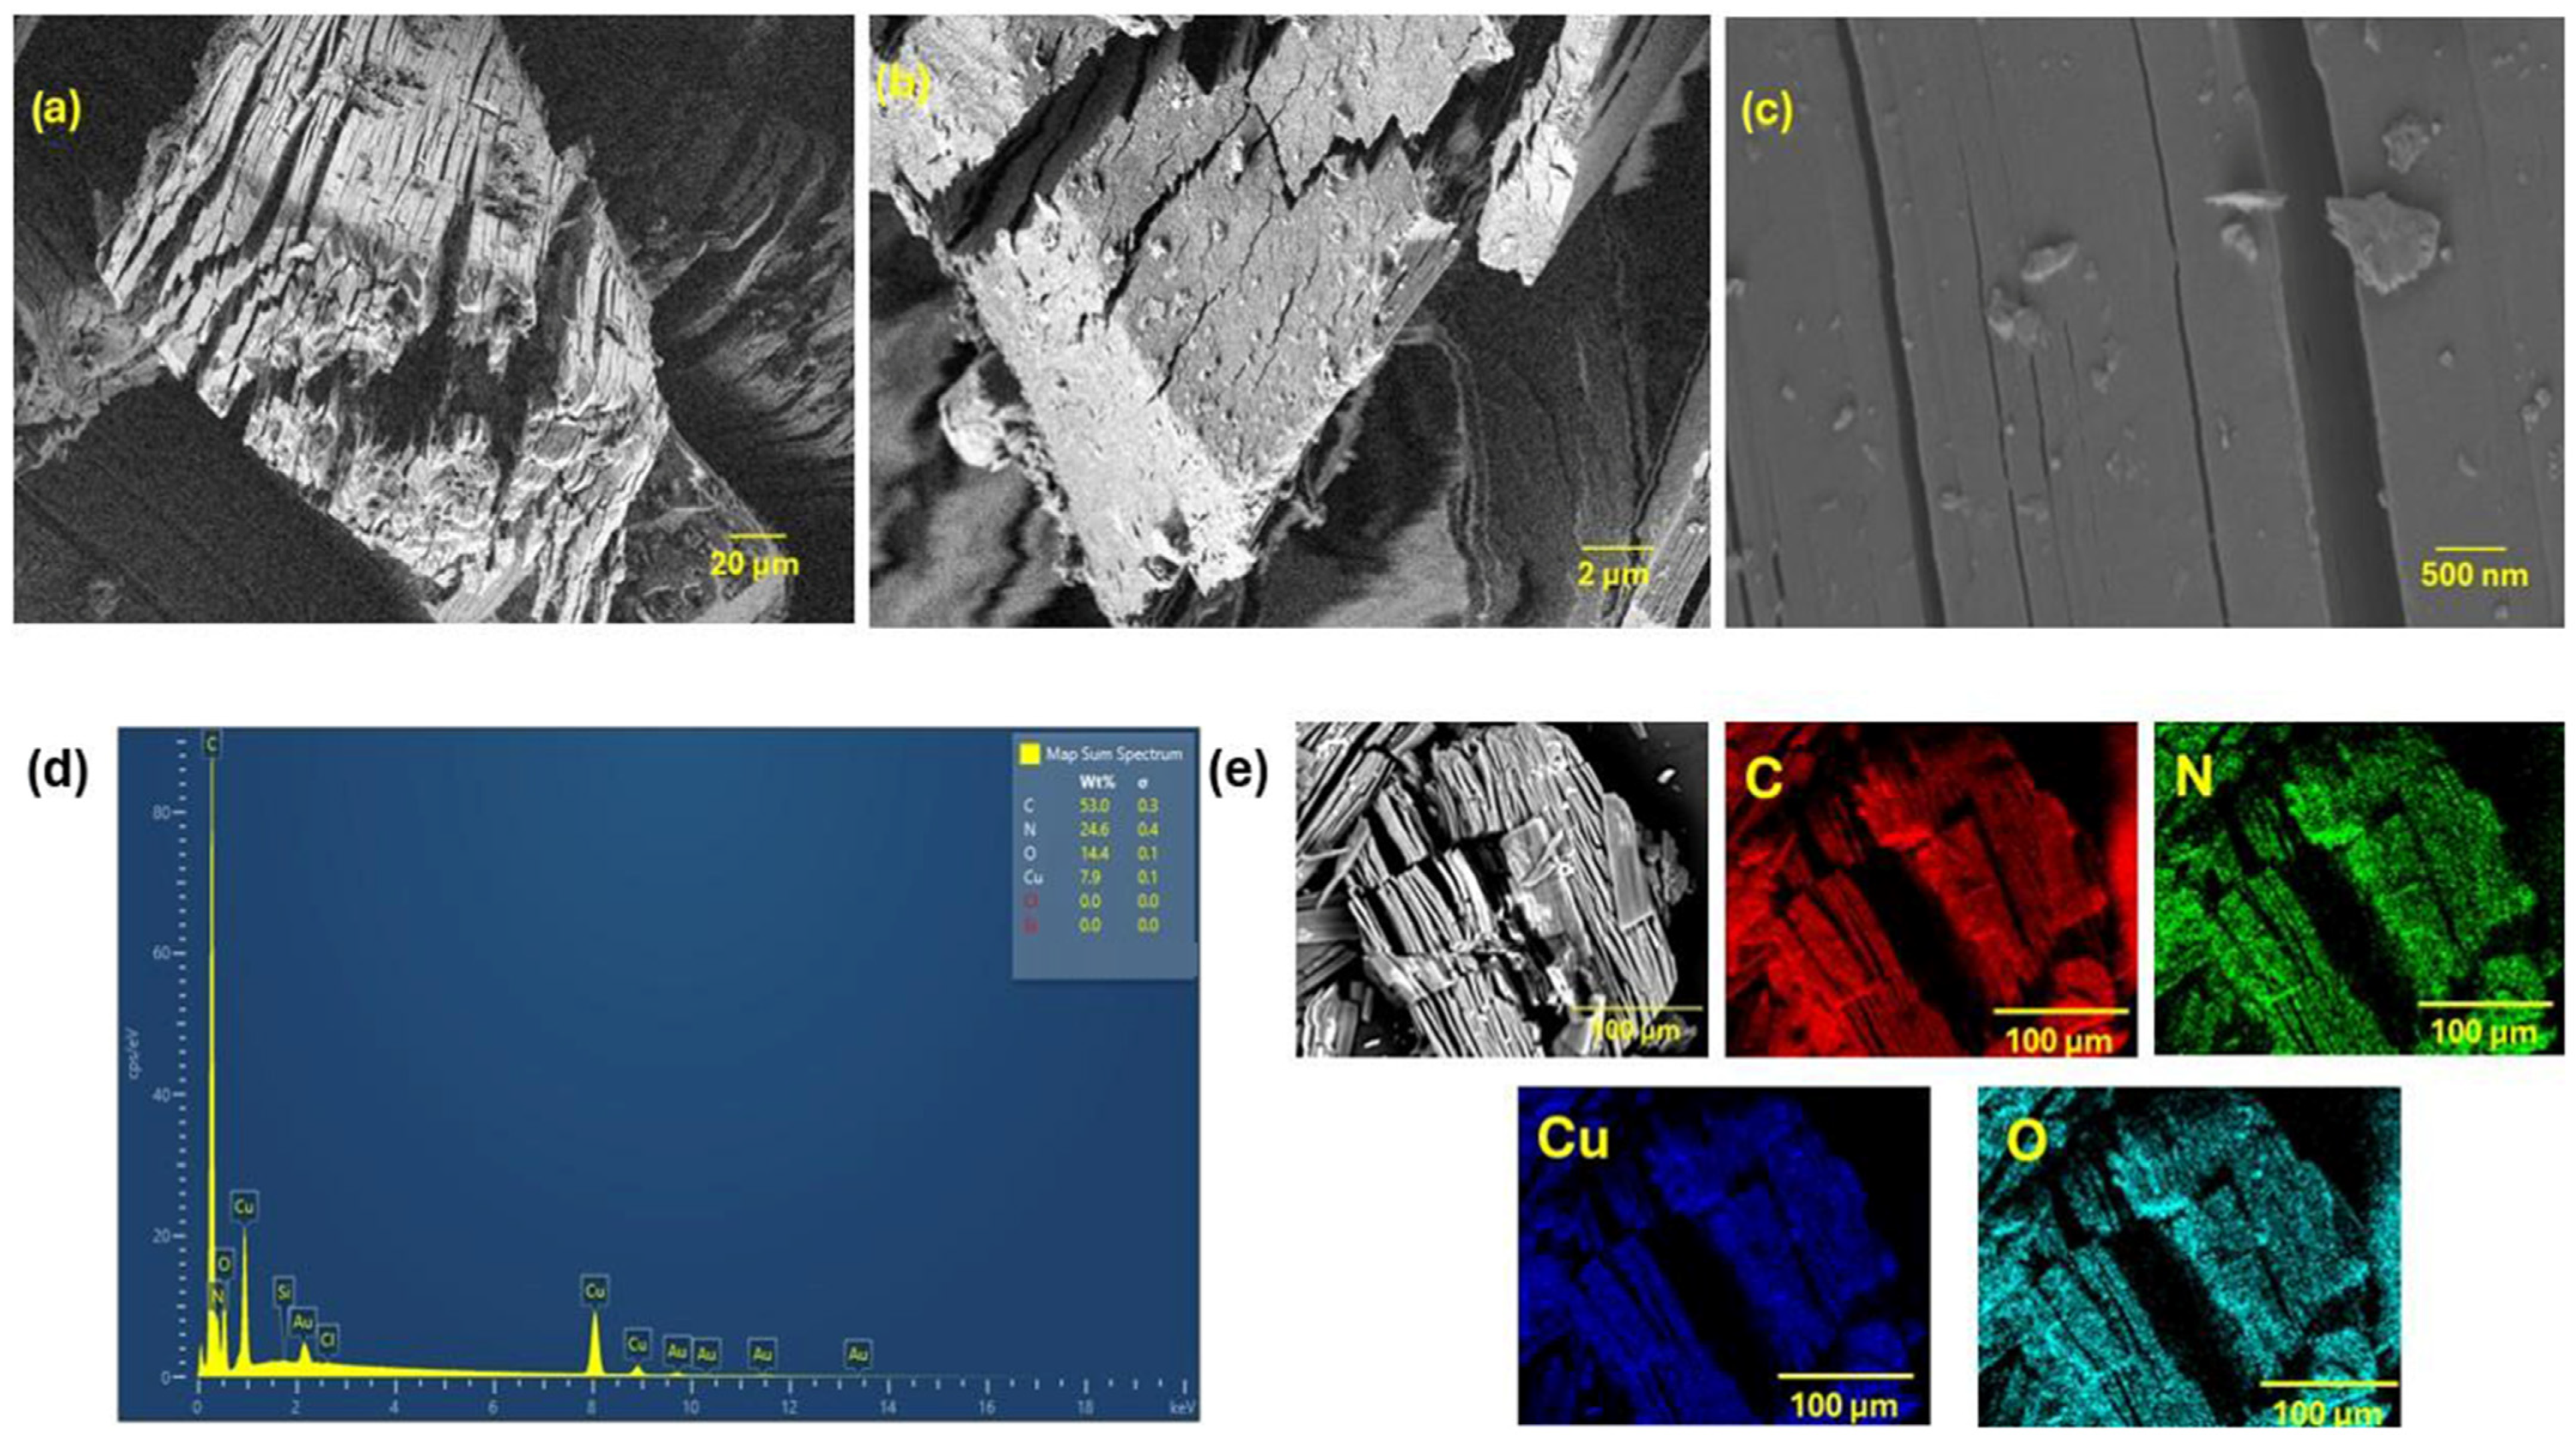
Figure 9 shows the morphological surface and elemental mapping of CCCu and CuHCF, respectively. Figure 9a shows the SEM and EDS of the CCCu compound. The material exhibits a distinct laminar architecture, composed of overlapping plates with sharp edges and broad, flat surfaces, features indicative of oriented crystalline growth. This layered geometry significantly enhances the surface-to-volume ratio, a critical parameter for interfacial processes such as adsorption or ion transport. At higher magnification (Figure 9b), microscale fissures and local porosity become evident, likely resulting from the release of crystallization water, in agreement with the endothermic signal observed in the DSC curve at ~80 °C.
These structural voids facilitate diffusion pathways and may influence electrochemical and thermal performance by enabling faster mass transport. In Figure 9c, the plates display fine striations aligned with the growth direction, confirming a highly regular, layer-by-layer assembly. Additionally, the presence of scattered nanoparticles suggests possible secondary nucleation centres that may act as active sites for electrochemical reactions or enhance conductivity and ion-exchange efficiency.
The global EDS spectrum (Figure 9d) confirms the expected elemental composition of the CCCu material, revealing weight percentages of C (53.0%), N (24.6%), O (14.4%), and Cu (7.9%), with no detectable impurities such as Cl or Si. Elemental distribution maps (Figure 9e) demonstrate a homogeneous spatial distribution of all elements. The uniform presence of Cu, without agglomeration, supports a well-integrated coordination environment within the organic matrix. These microstructural and compositional features validate the empirical formula of CCCu and correlate with the thermal degradation behaviour described previously. The material’s high surface area, laminar morphology, and elemental purity form a consistent basis for its potential applications in electrocatalysis and corrosion-resistant systems.
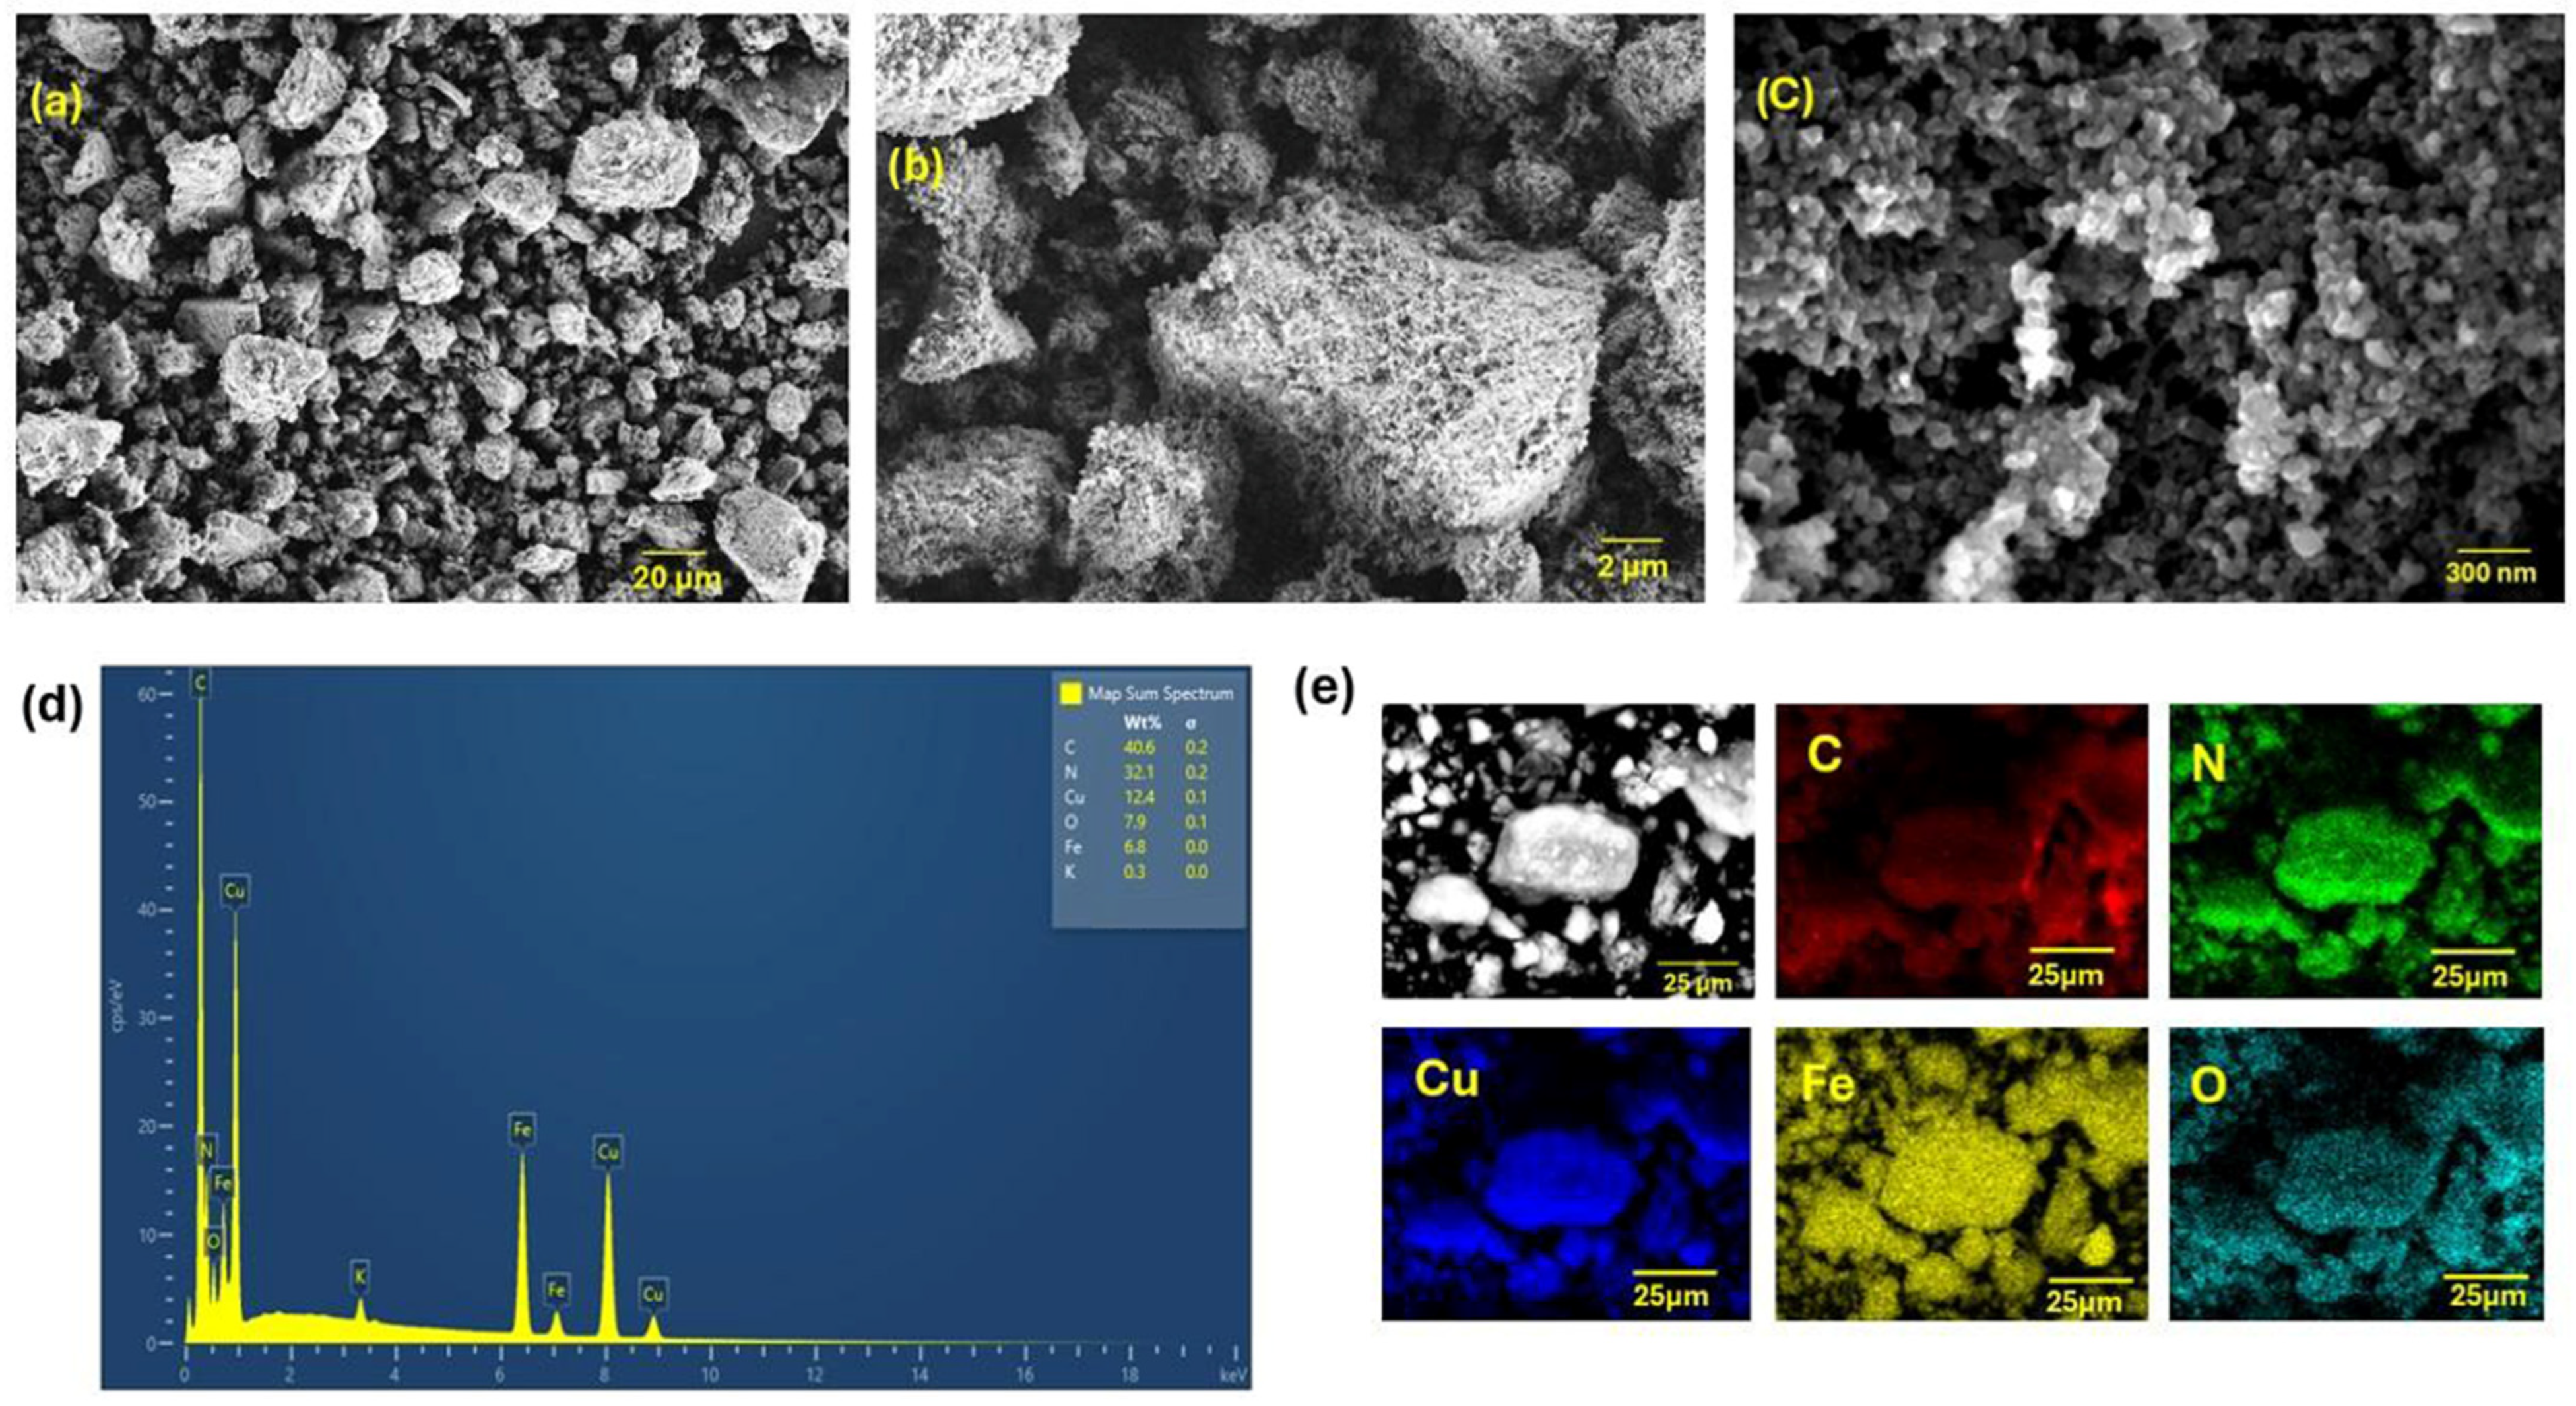
In contrast, Figure 10 presents the morphological and compositional analysis of the CuHCF compound. In Figure 10a, the micrographs reveal a spherical particle morphology, with diameters ranging from 5 to 30 μm and a relatively uniform dispersion. This sub-micrometric granular architecture is beneficial for electrochemical applications, offering a balanced compromise between surface area and packing density.
Upon magnification (Figure 10b), the surfaces appear rough and porous, containing both external voids and internal nodules, which likely correspond to zones of localized crystallization. This heterogeneous texture enhances the reactive surface area and facilitates electrolyte diffusion, both key factors for improving electrochemical efficiency. At the nanoscale (Figure 10c), the images confirm a hierarchical structure: aggregates of nanoplatelets (50–200 nm) form mesoporous spherical assemblies without observable microcracks. This structural motif not only ensures mechanical robustness under thermal cycling but also promotes rapid ion transport, essential for long-term operational stability.
The EDS spectrum (Figure 10d) provides valuable insights into the elemental composition and offers a preliminary indication of the material’s stoichiometry. The results align with the expected composition of the Prussian Blue analogue [Cu2+][Fe2+(CN)6]·H2O. The detected elements and their corresponding weight percentages are C (40.6%), N (32.1%), O (12.4%), Cu (7.9%), Fe (6.8%), and K (0.3%). The presence of potassium suggests its partial incorporation into the lattice or occupation of interstitial sites. Elemental mapping (Figure 10e) reveals a homogeneous distribution of the key framework elements (C, N, Cu, Fe) and hydration-associated oxygen, while potassium appears scattered yet localized, supporting its assignment to interstitial positions. Taken together, these morphological and compositional insights reinforce the interpretation of TG/DSC data, particularly the initial mass loss associated with water desorption and the stability of the cyanide-metal network beyond 250 °C. Overall, the mesoporous architecture, nanoscale organization, and chemical homogeneity of CuHCF make it a robust candidate for catalysis and seawater electrolysis under harsh conditions.

3.5. Corrosion Analysis

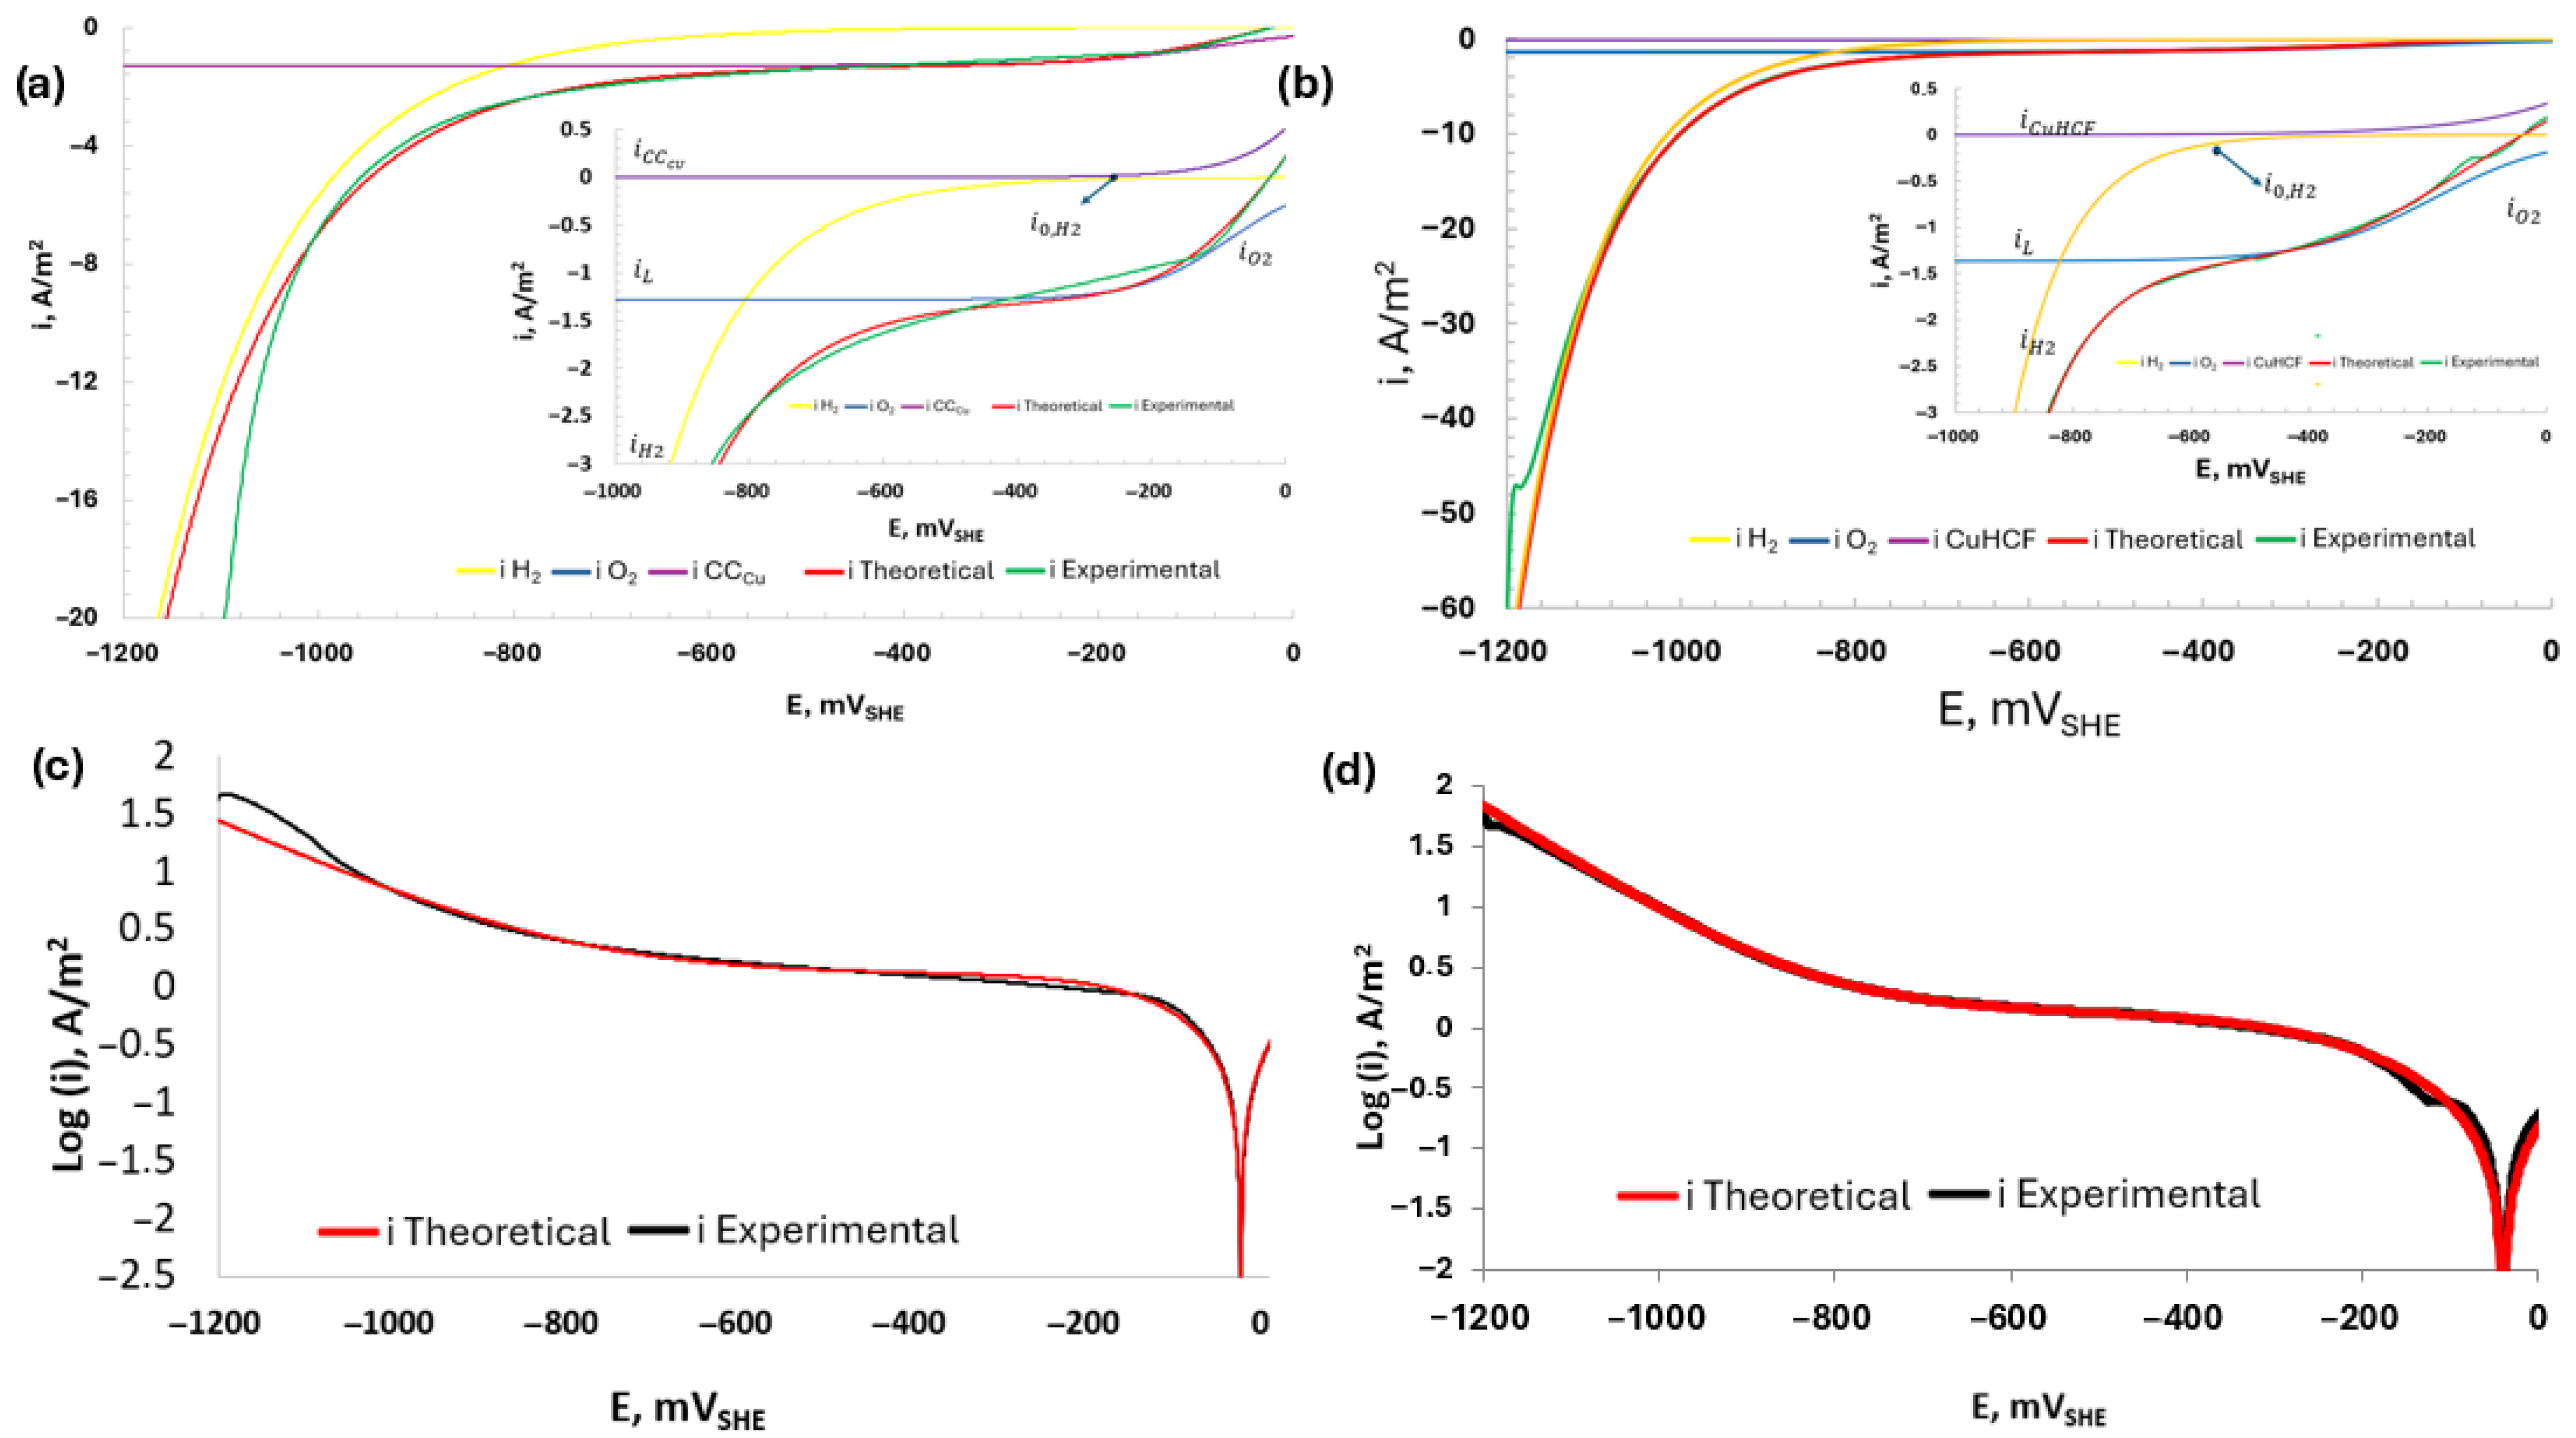
The novelty of this section is demonstrating for the first time that coordination compounds traditionally employed in electrochemical energy storage can act as active electrocatalysts in 0.5 M NaCl media for green H2 production. This discovery redefines their application field, opening a new paradigm centred on hybrid materials and offering a viable alternative for the direct electrolysis of seawater and other saline solutions, which represents a critical challenge for the energy transition. However, a long-term study demonstrating the stability of corrosion-resistant material using chronoamperometry was not conducted, which is a limitation of the manuscript. The kinetic results were obtained by fitting the data of the experimental potentiodynamic polarization curves with the superposition model based on mixed-potential theory, as shown in Figure 11. This presents a comparison between the experimental results and those obtained through the superposition model based on mixed potential theory for the CCCu and CuHCF in saline media. Figure 11a,b shows how the model accurately reproduces the overall electrochemical response, allowing the partial contributions of the HER, ORR, and the oxidation processes of the material itself to be deconvoluted. The Tafel plots shown in Figure 11c,d confirm this consistency, demonstrating a good correlation between theoretical and experimental data across the entire potential range. These results validate the proposed approach to describe the complexity of systems in neutral chloride solutions and provide a solid basis for comparing the catalytic activity and corrosion resistance of both materials.
This approach allows simultaneous integration of charge transfer processes, mass diffusion limitations, and oxidation mechanisms, providing a comprehensive description of the electrochemical behaviour of coordination compounds in neutral saline solutions. The following metals (i) A36, (ii) AISI1020, (iii) A630-420H steels [39], (iv) Ti [40,56,57], with copper-based coordination compounds such as (i) CuHCF and (ii) CCCu, were tested in aerated 0.5 M NaCl solution. The results reveal fundamental differences in corrosion stability and electrocatalytic activity. For carbon steels, the i c o r r are in the range of 1.36–3.02 A/m2 and E c o r r between −221 to −320 mVSHE, respectively. However, since these materials are unconventional for the HER (i.e., molecular or coordination-based, non-noble materials directly evaluated in neutral chloride media), there is currently no systematic literature supporting their use as HER electrocatalysts under such conditions. Triazole-based coordination compounds and Prussian Blue Analogue-type materials remain largely unexplored for HER in aerated NaCl electrolytes. These values reflect active dissolution performance in the presence of Cl− ions, resulting in limited resistance in saline environments. Ti, in contrast, presents a more negative potential (–456 mVSHE) due to the spontaneous formation of a protective TiO2 film, with an i c o r r of 0.019 A/m62. With respect to the HER, steels display exchange current densities (|i0,H2 |) in the range of 0.02 to 0.39 A/m2, while Ti exhibits even lower values in the range 0.001 to 0.01 A/m2, remaining to its surface passivation by TiO2, confirming its poor catalytic efficiency in saline solutions. In contrast, the coordination compounds CuHCF and CCCu show evidently superior performance, with |i0,H2| values of 0.019 A/m2 and 0.062 A/m2, respectively. It is important to note that the exchange current density values are still lower than those obtained on Pt, Ir, or Re electrodes; however, the values are higher than those obtained on Cu, Ni, Co, and Ag electrodes, which represents comparative performance. This translates into a much more pronounced catalytic functionality. The steels exhibit HER tafel slopes |bH2| of 196 to 270 mV/dec and ORR tafel slopes |bO2| of 227 to 247 mV/dec, whereas the coordination compounds record 301 mV/dec (CCCu) and 222 mV/dec (CuHCF) for HER, along with ORR slopes of 157 to 238 mV/dec, respectively. The Tafel slope and exchange current density for hydrogen evolution on carbon steel exhibit considerably greater variability than those measured on stainless steel. Although the exchange current densities are of comparable magnitude, the Tafel slopes |bH2| for carbon steel typically range from 240 to 330 mV/dec. This pronounced variation in the kinetic parameters of hydrogen evolution on carbon steel has also been reported by other authors, even when measured in non-buffered solutions. For the other way, NiMoP coatings, the reported |bH2| range between 101 and 275 mV/dec, in alkaline media, in comparison, CuHCF exhibits a value of 222 mV/dec. This indicates that, although copper coordination compounds display appreciable semi-electrocatalytic activity, in terms of hydrogen evolution kinetics [58,59]. Their electronic conductivity, ion transport, and interfacial charge-transfer mechanisms differ, leading to large kinetic overpotentials. Although these results diverge from classical expectations, they can be explained by the superposition model for kinetic and diffusional phenomena. Relevant factors include (i) mass transport limitations caused by the low solubility of oxygen in saline solutions and the sluggish kinetics of proton adsorption in neutral media; (ii) simultaneous contributions from metal corrosion processes and cathodic reactions, which distort the linearity of Tafel plots; and (iii) surface heterogeneity, arising from corrosion roughness in steels and the pseudocapacitive microstructure of CuHCF; modifying activation energies for HER and ORR; and (iv) the intrinsic use of the superposition model, which integrates multiple reactions into a single fit, thereby altering classical mechanistic interpretation. In this sense, rather than being considered an experimental anomaly, the elevated Tafel slopes reflect the real complexity of electrochemical processes in neutral saline systems, which differ significantly from those in acidic or alkaline environments. The most relevant point is that, even under such hostile conditions, CuHCF achieves a Tafel slope |bH2| of 222 mV/dec comparable to or even lower than advanced systems such as NiMoP [60], indicating a reduced overpotential requirement for H2 generation.
Concerning the ORR, both coordination compounds exhibit low exchange current densities (see Table 1), consistent with non-noble materials. Nevertheless, CuHCF outperforms CCCu, showing a higher limiting current is −1.362 A/m2 vs. −1.279 A/m2, attributable to its highly ordered pseudocapacitive structure that facilitates ionic and electronic transport. On the anodic side, CCCu presents a lower material oxidation exchange current (i0, M = 0.395 A/m2) compared with CuHCF (i0, M = 0.740 A/m2), suggesting slightly greater resistance to electrochemical dissolution. Overall, the comparison between metals and coordination compounds highlights a key dichotomy: while metals, particularly Ti, ensure durability due to their corrosion resistance, they lack significant catalytic activity in saline environments. Conversely, coordination compounds sacrifice stability but deliver unprecedented electrocatalytic functionality in neutral chloride media. This shifts the kinetic and electrochemical parameters of HER and ORR, with Tafel slopes that reveal more favourable kinetics than expected, thus positioning these compounds as unconventional yet promising candidates for seawater electrolysis and sustainable hydrogen production. Table 1 presents the kinetic and corrosion parameters for CCCu and CuHCF immersed in 0.5 M NaCl.
Transition-metal phosphides such as Ni2P, CoP and FeP [61,62], typically exhibit |bH2| values between 80 and 140 mV dec−1 and overpotentials of 180–250 mV at 10 mA cm−2 but suffer from severe surface oxidation and chloride-ion-induced corrosion during prolonged electrolysis, which reduces their operational activity to around 10 h. Similarly, transition-metal sulphides such as MoS2, Ni3S2 and CoSx achieve comparable HER kinetics (|bH2| of 120–180 mV dec−1) but undergo progressive sulphur leaching in media with high Cl− concentration. In contrast, the coordination-based compounds studied here (CCCu and CuHCF) display higher |bH2| values ranging from 222 to 301 mV dec −1, associated with their semi-electrocatalytic behaviour; nevertheless, they retain electrochemical activity in neutral saline electrolytes, where no evidence of pitting or passivation was observed, demonstrating their intrinsically corrosion-resistant character [63,64,65].

3.6. DFT Study of the Hydrogen Evolution Mechanism and Crystalline Stability

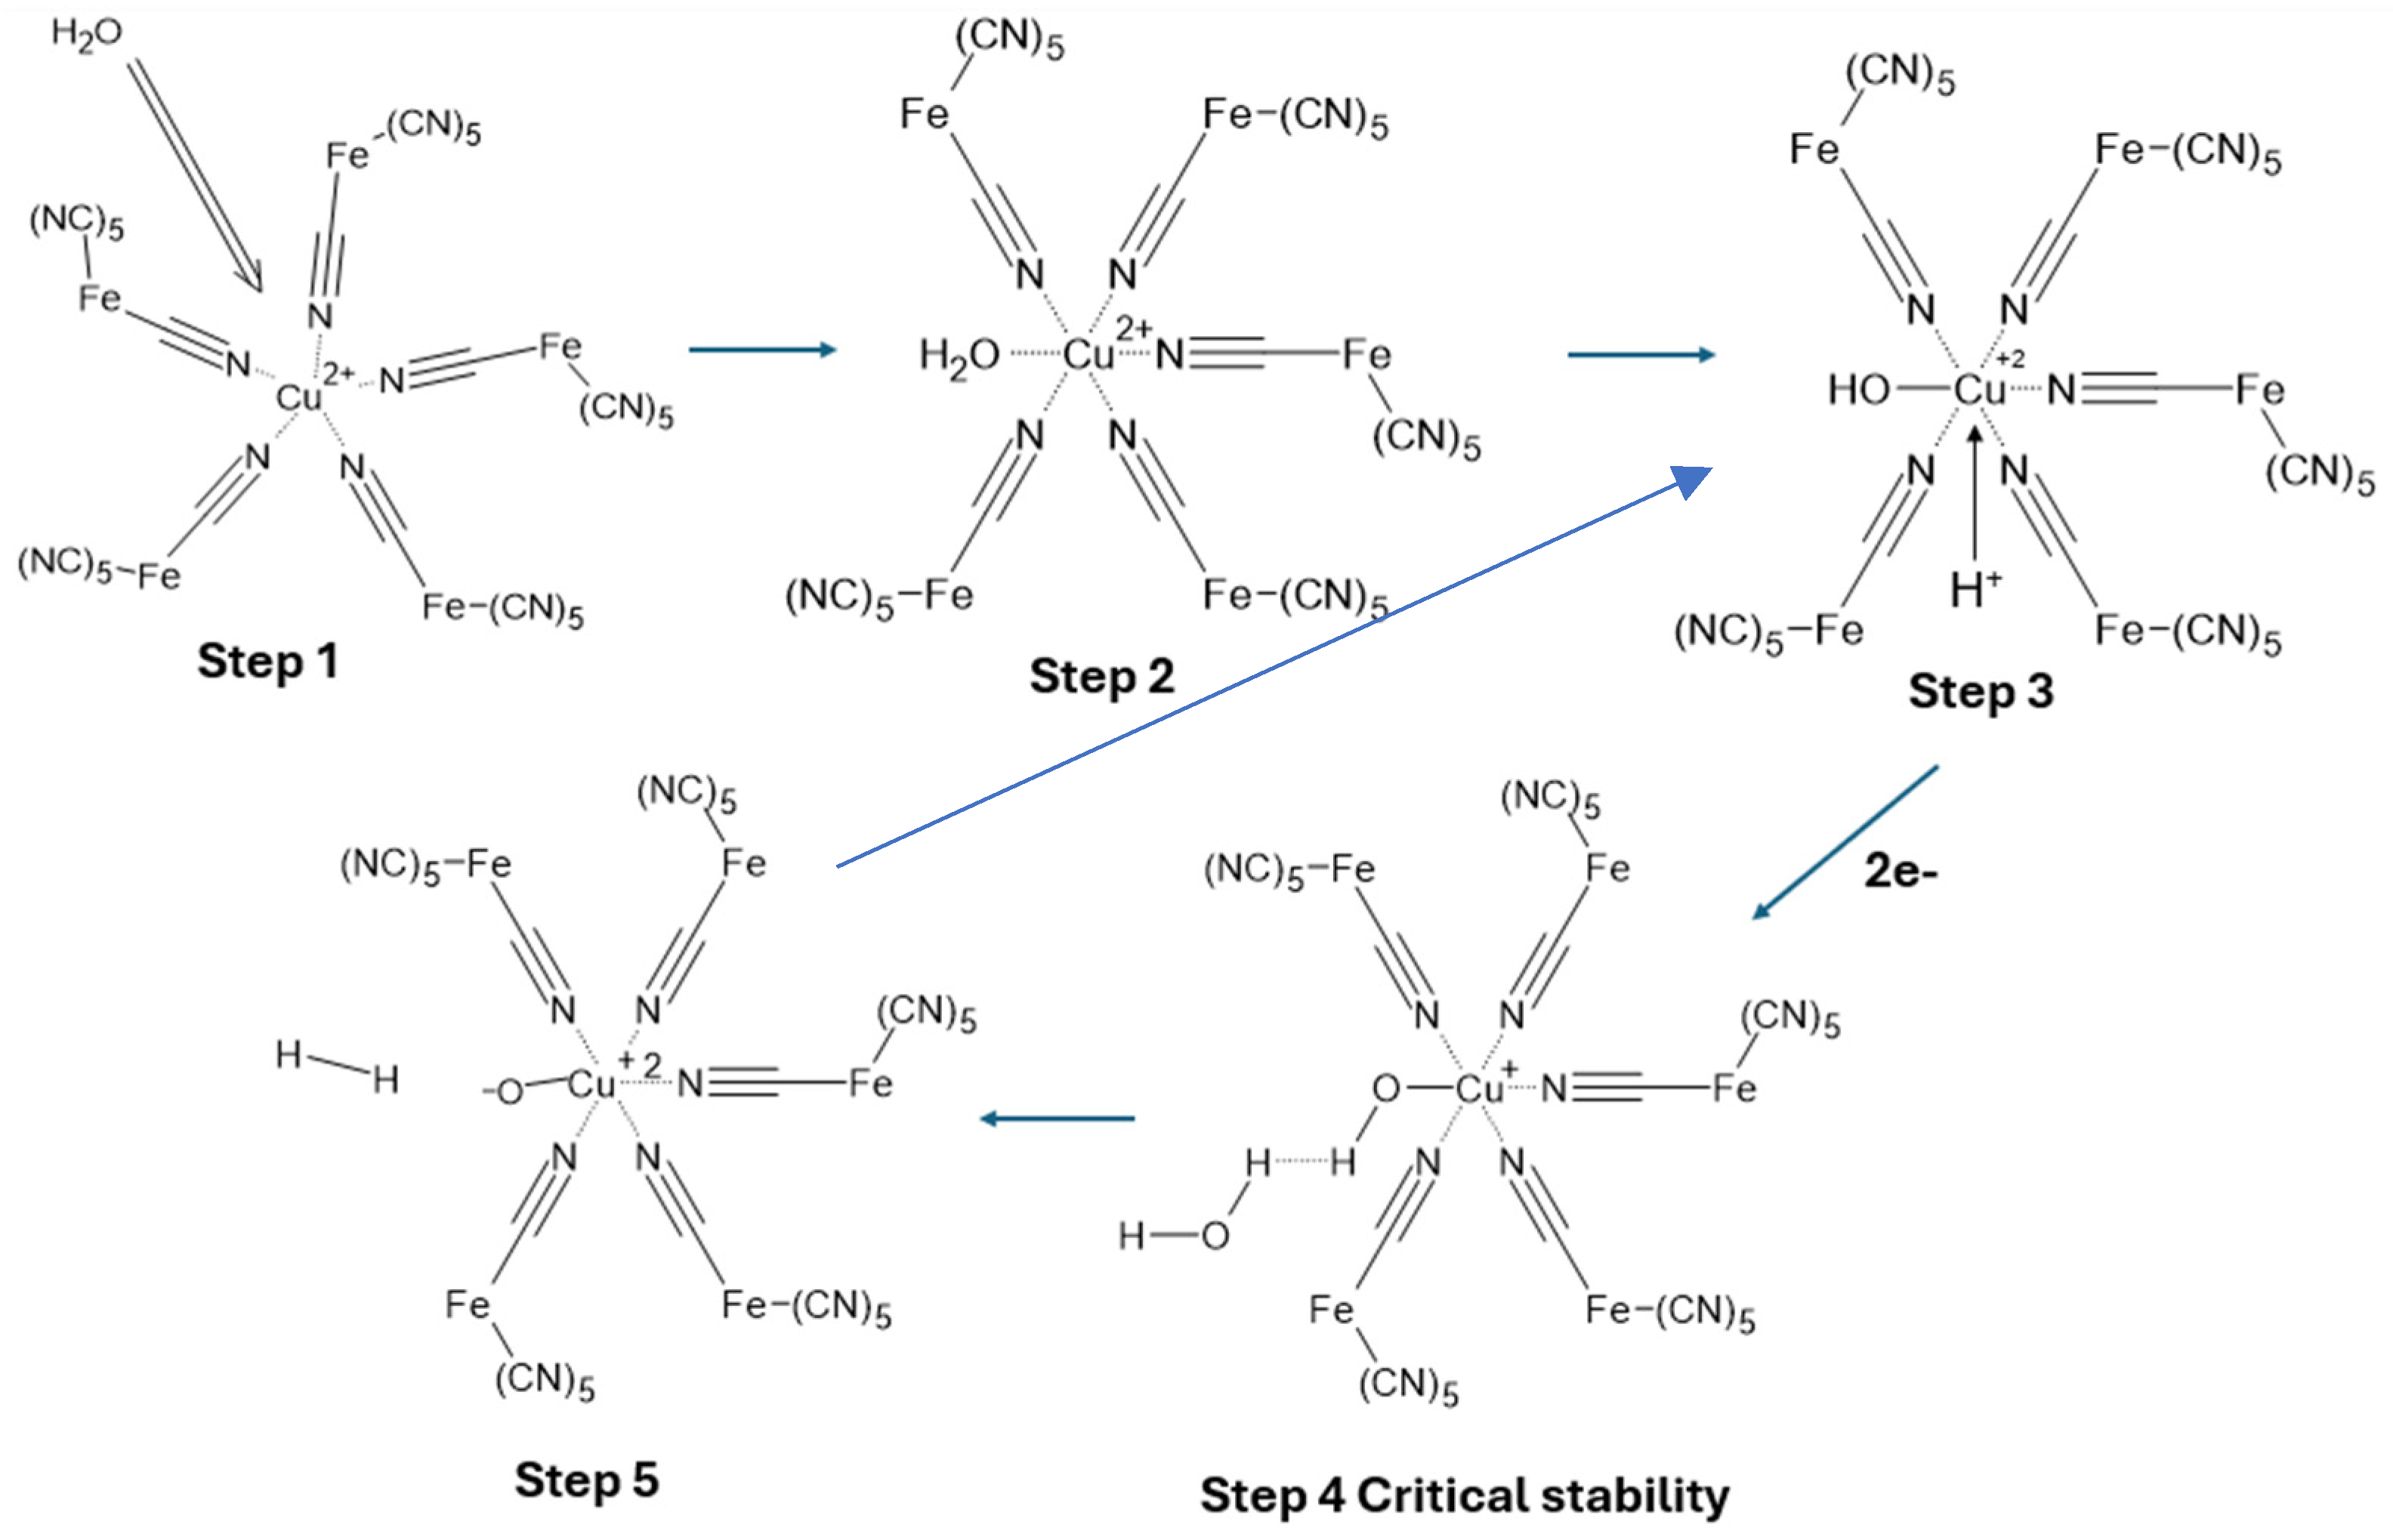
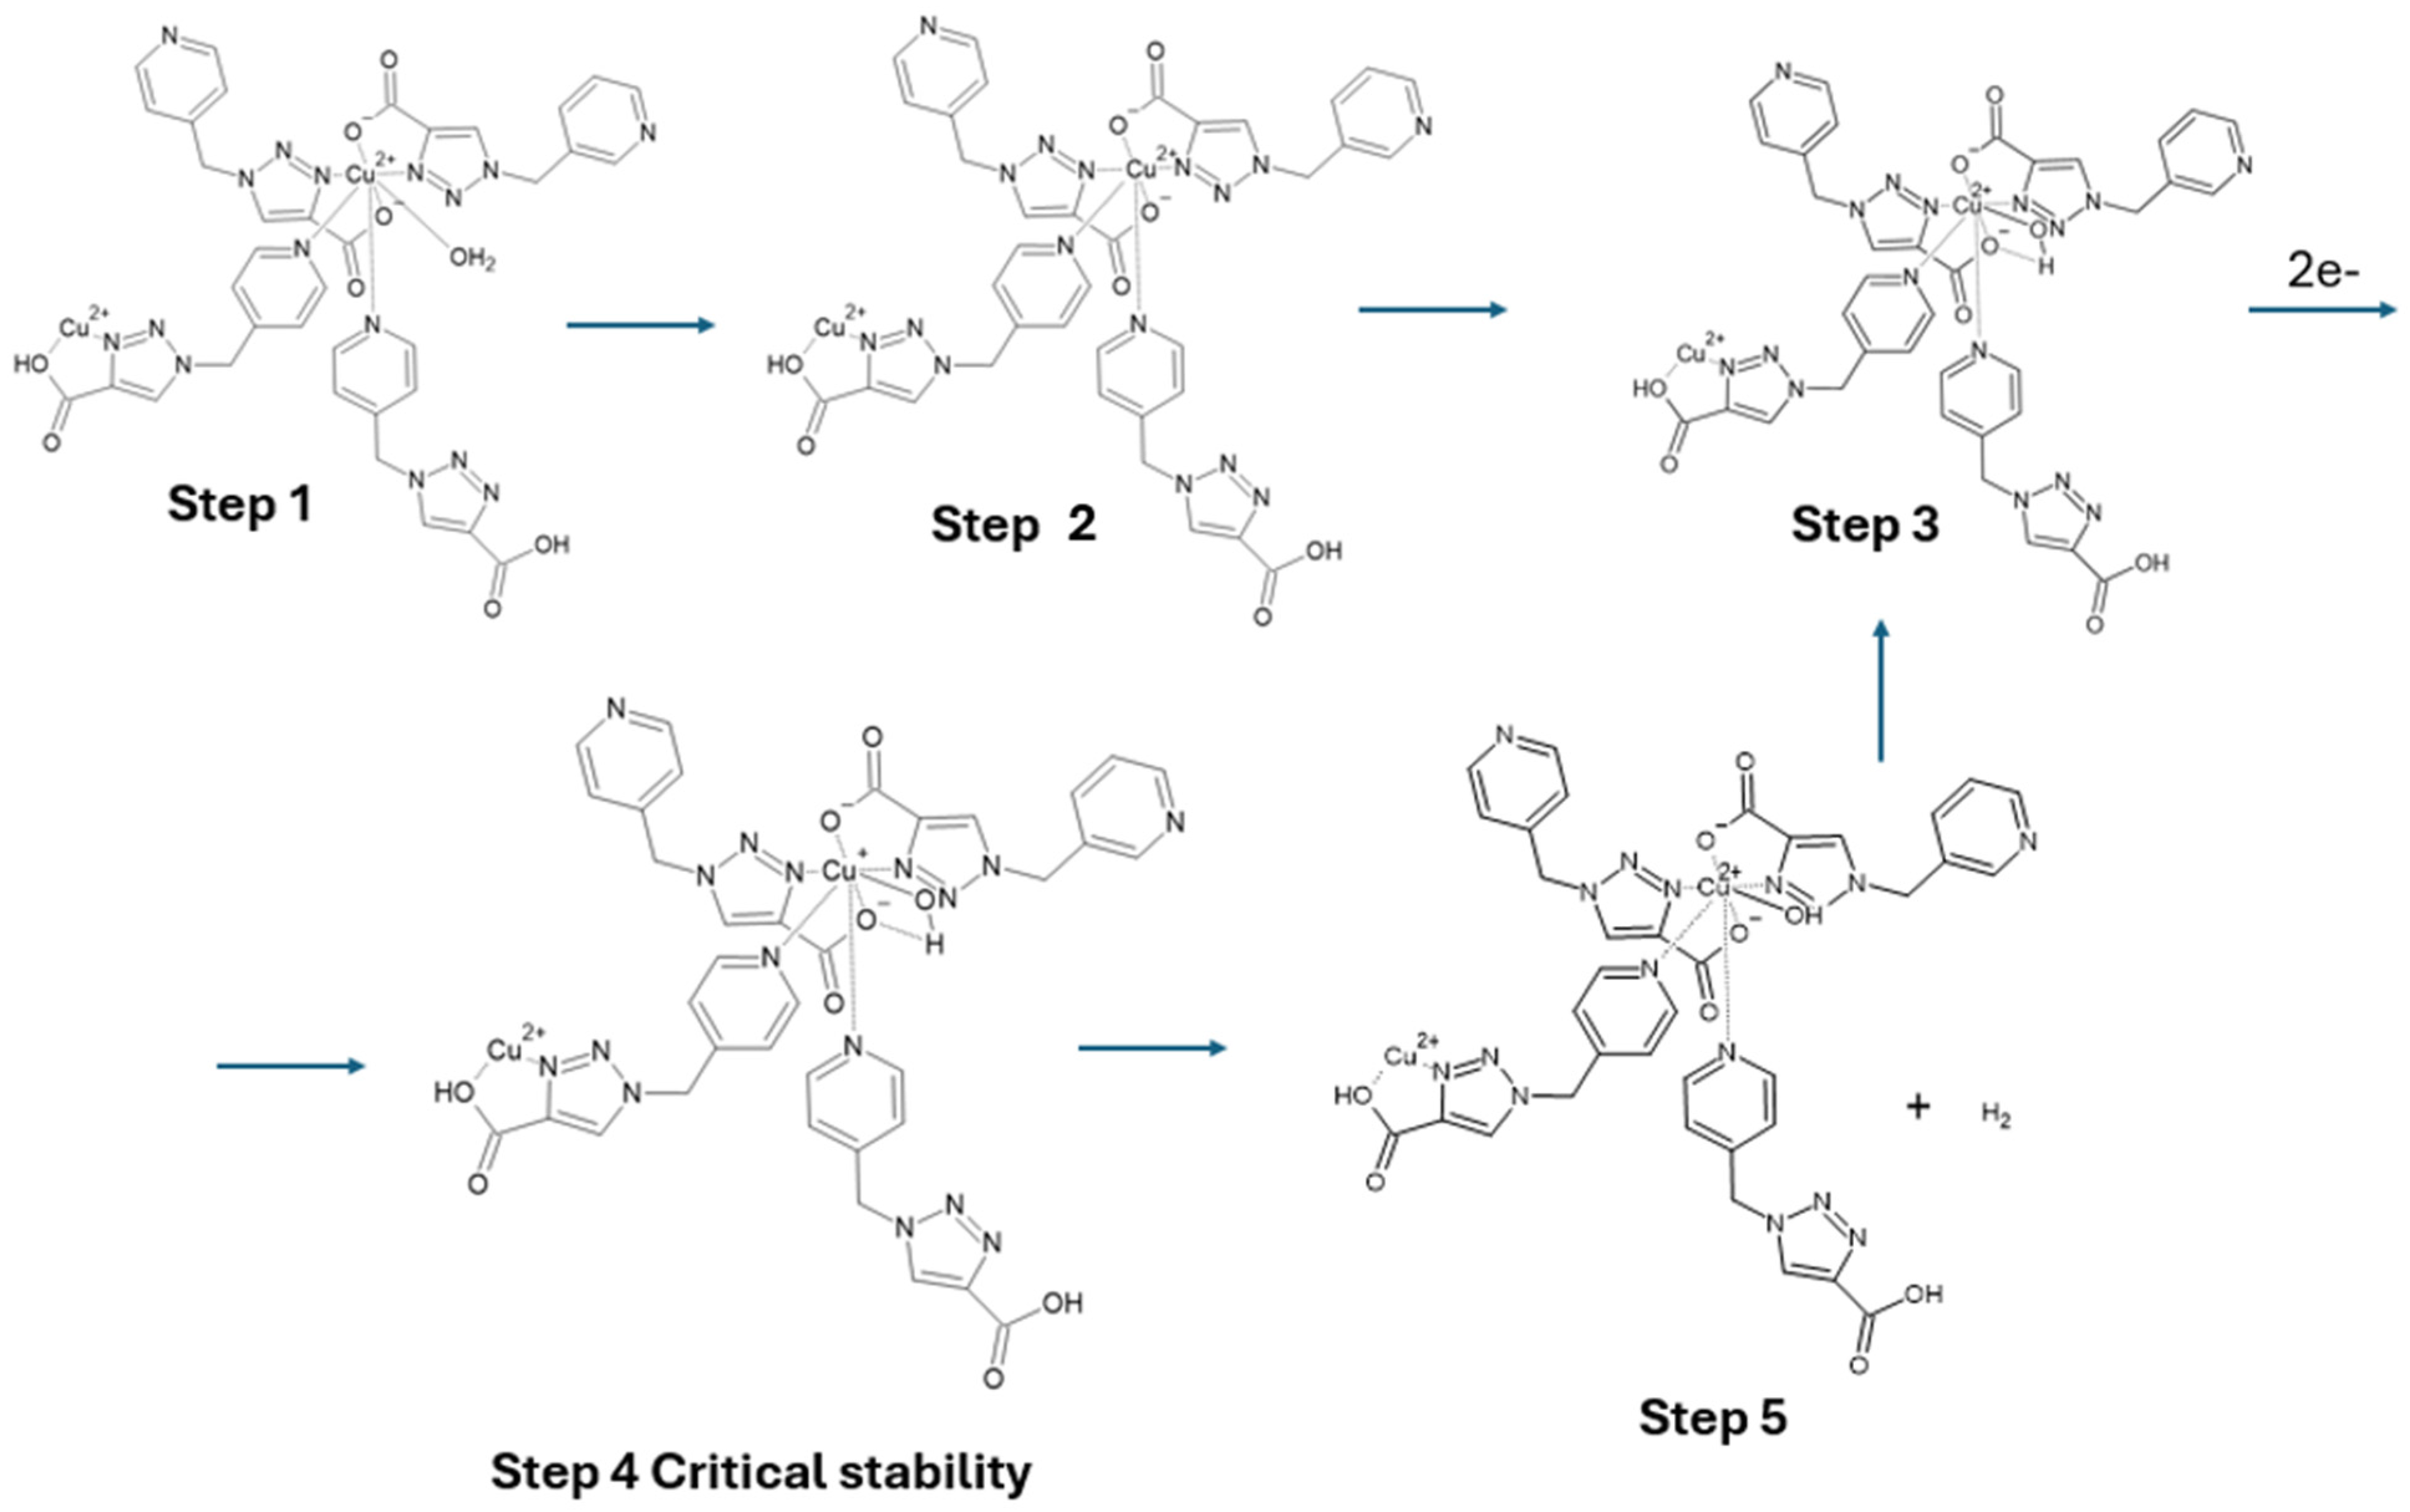
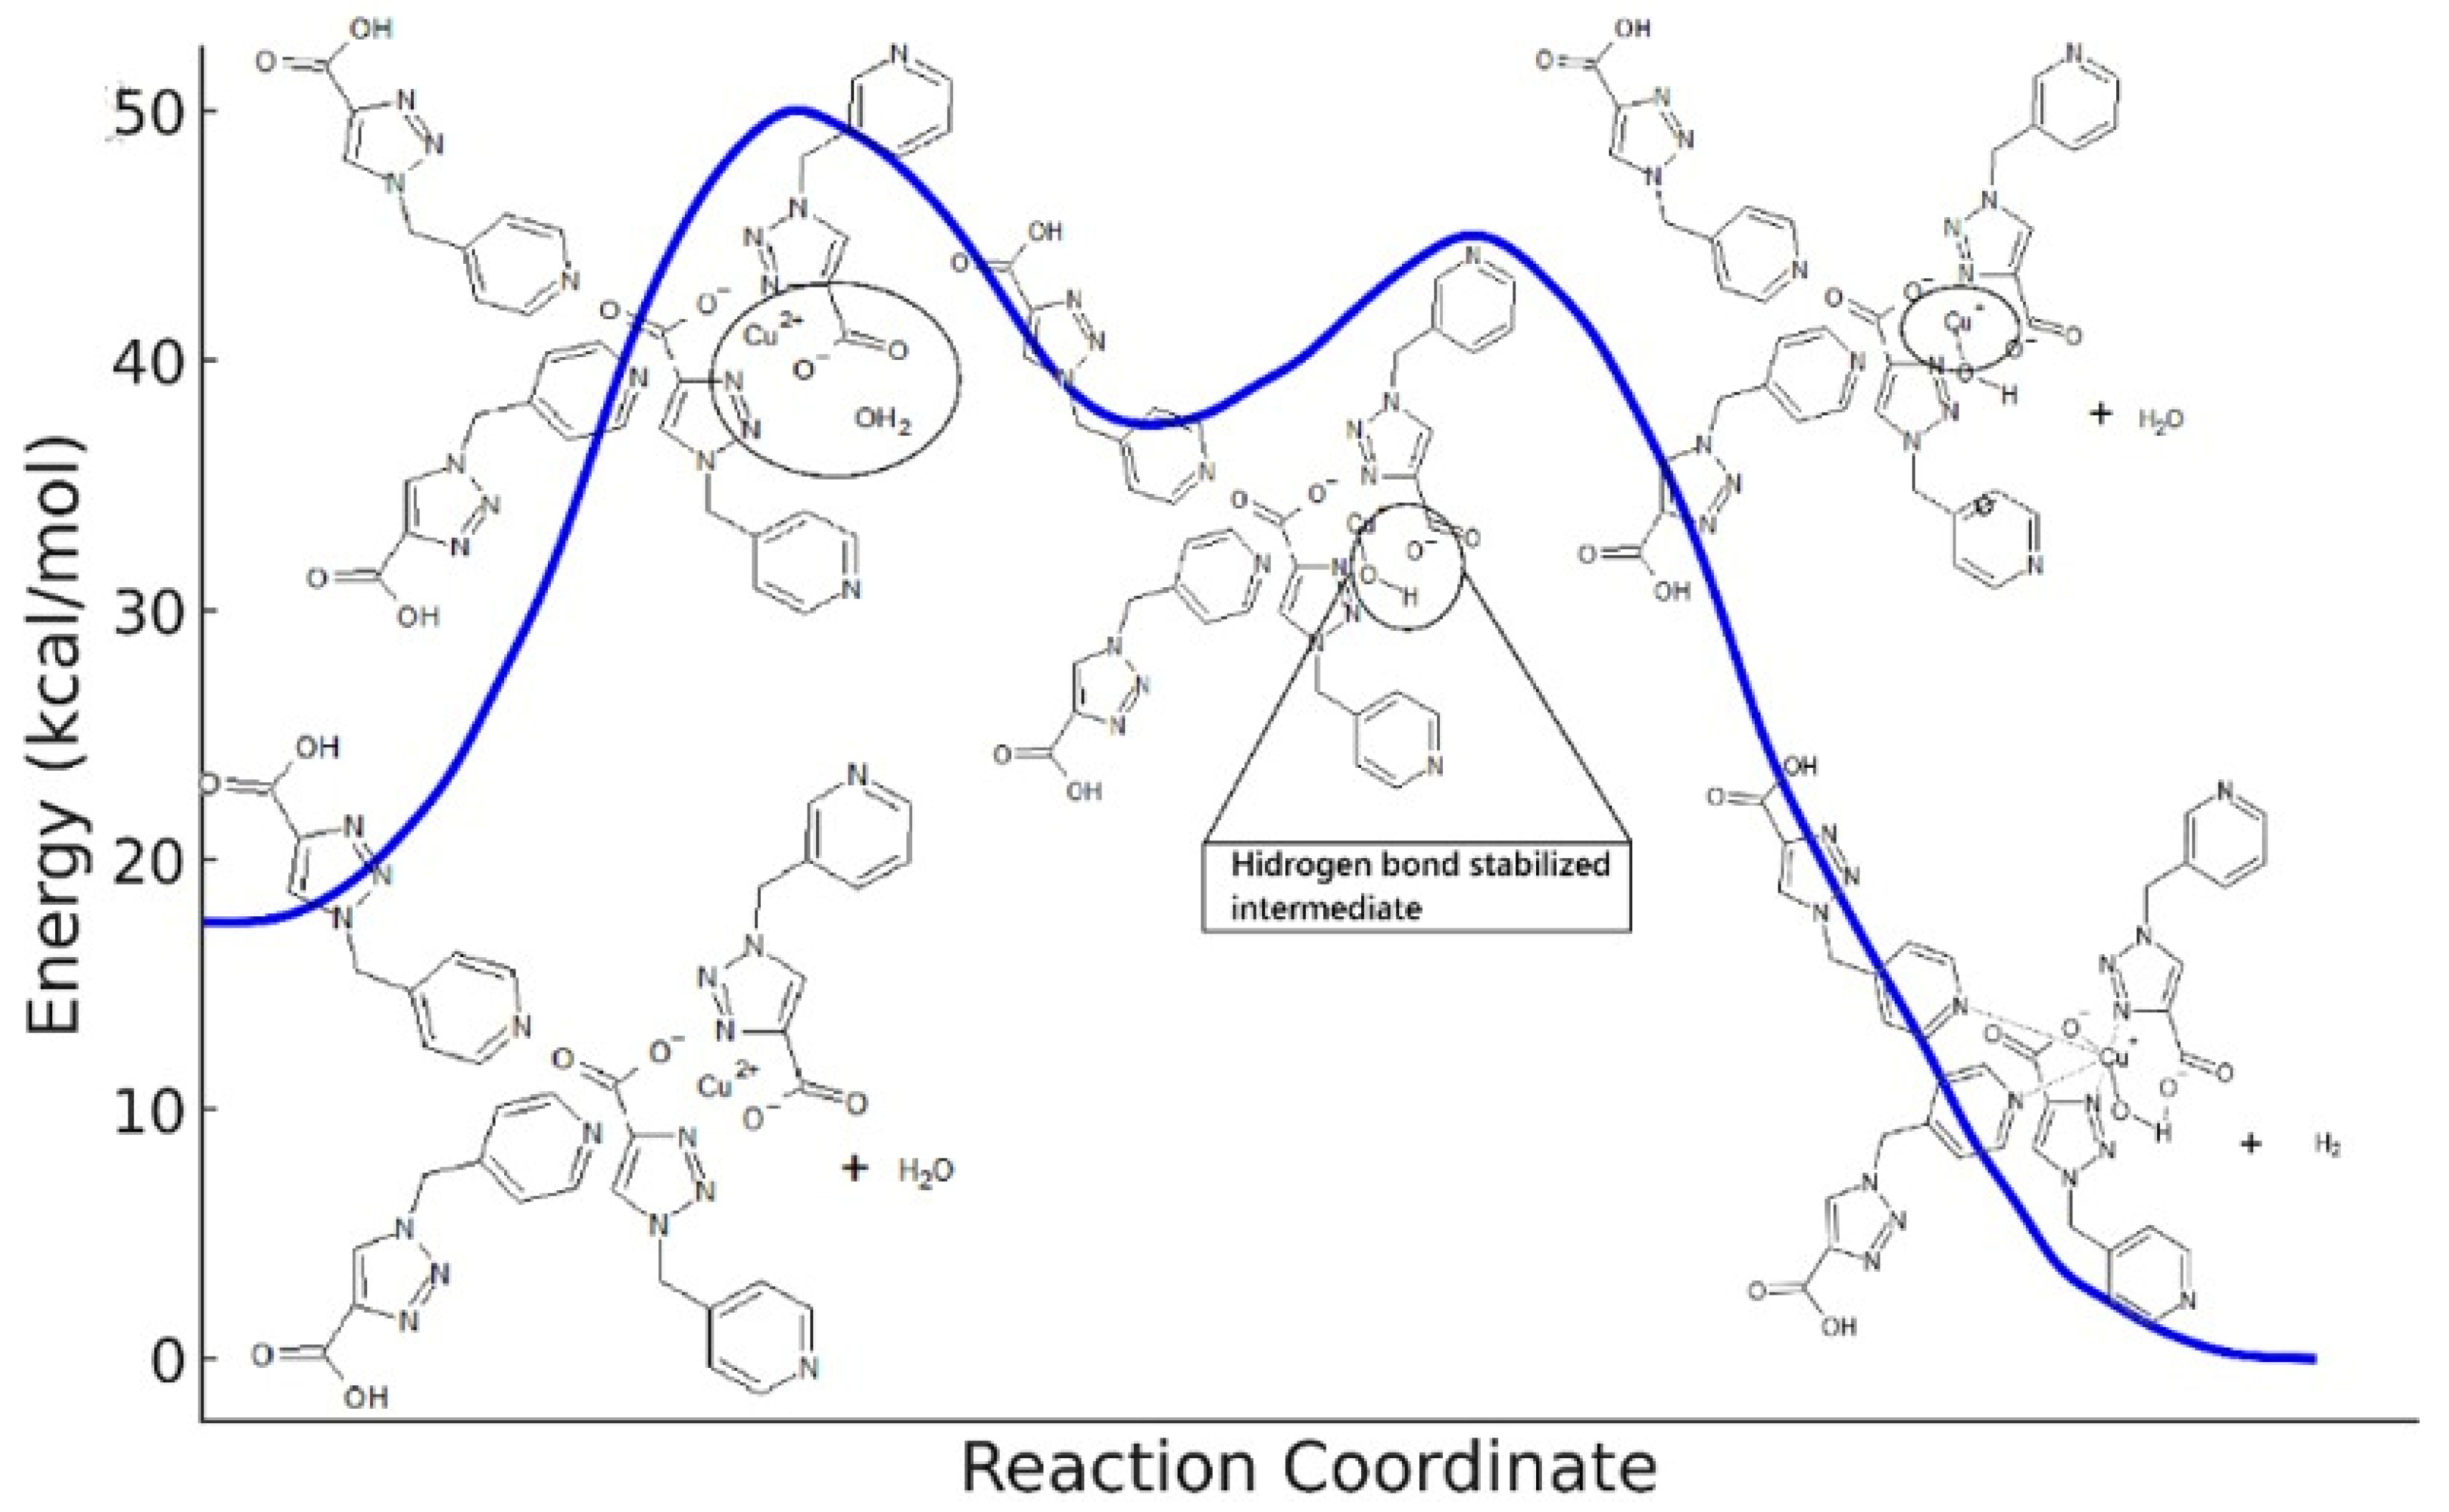
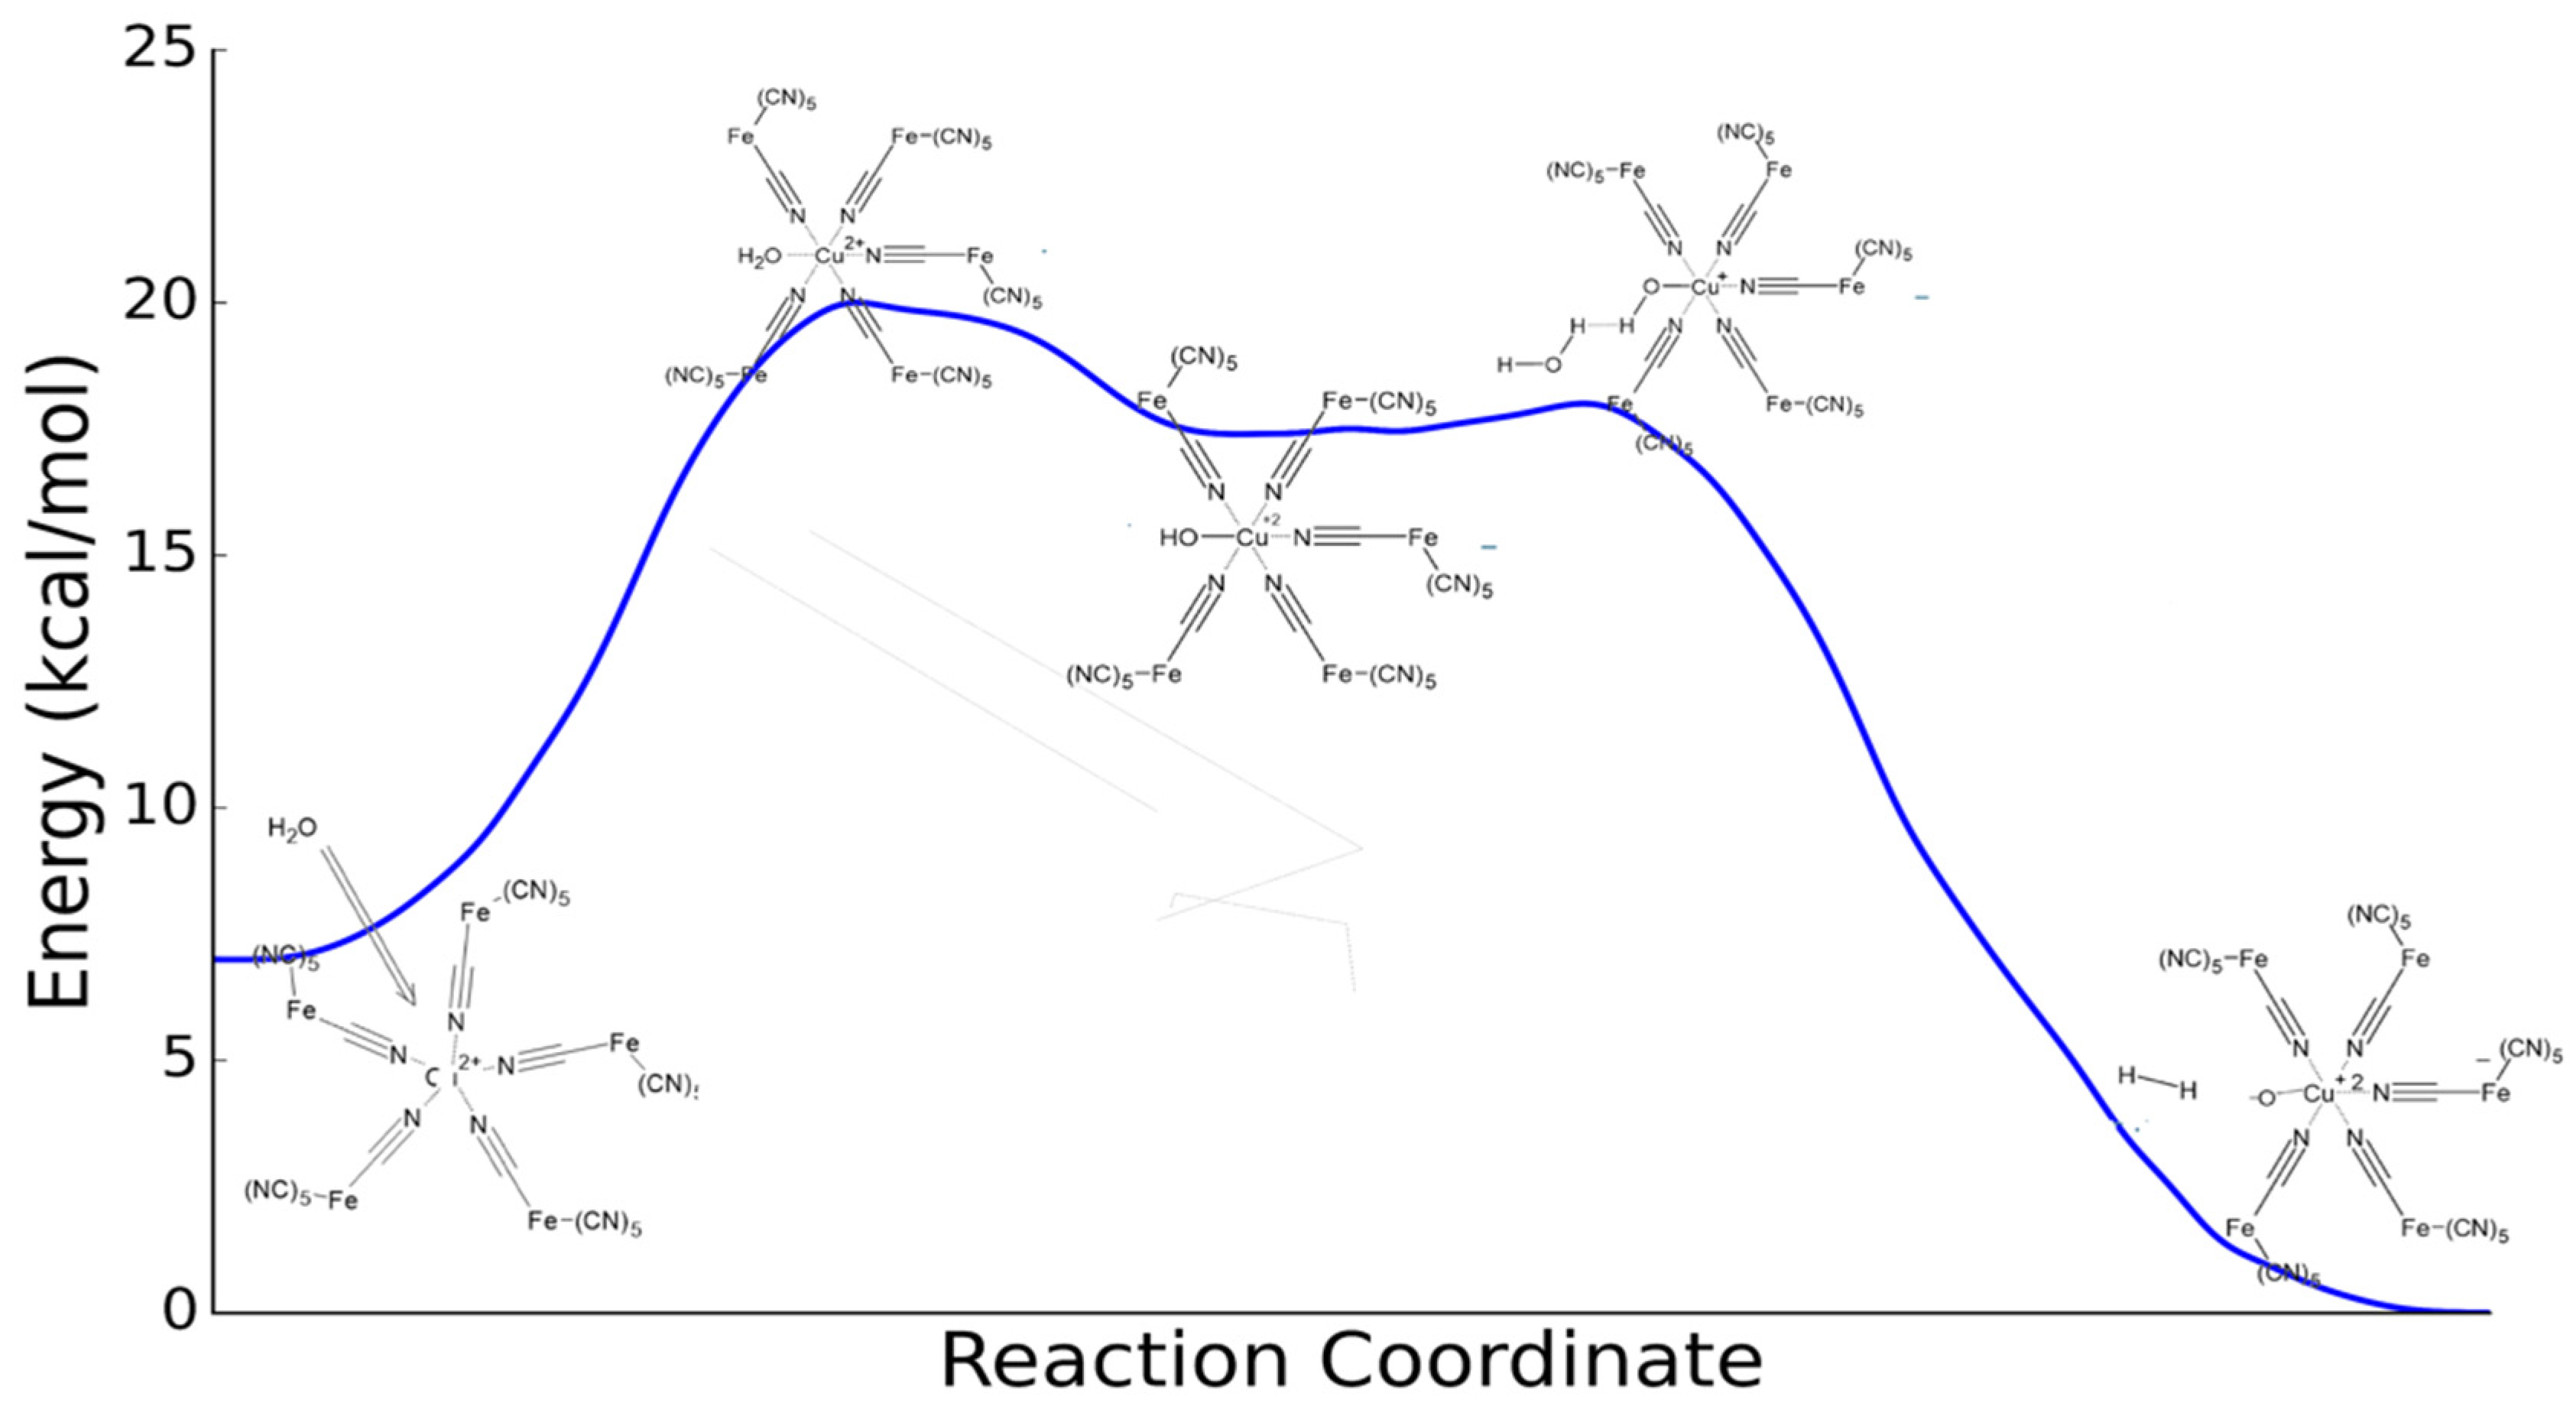
Theoretical chemistry makes it possible to analyze phenomena that, to date, cannot be directly measured by experimental techniques. In this context, we present a theoretical study aimed at validating a possible mechanism for the HER, with the objective of providing a detailed and coherent description of the catalytic process in agreement with the experimental data. In this way, the electronic structure of the system can be correlated with its geometry and with the results observed experimentally. Figure 12 illustrates the proposed reaction mechanism, which consists of five steps. The first two steps correspond to the activation stage, while steps 3, 4, and 5 represent the catalytic cycle. Thus, once activation has taken place (steps 1 and 2), the catalytic process involves only steps 3–5. After water enters the crystal, it coordinates to the copper center and stabilizes as a hydroxide ion strongly bound to the metal. This generates an intermediate reaction located in the valley between transition state 1 and transition state 2 (see Figure 12). Upon electron injection, the copper center undergoes reduction from Cu2+ to Cu+, enabling the formation of a hydride species which, upon coupling with a proton or water molecule, evolves into gaseous hydrogen (Figure 12, Figure 13 and Figure 14). Subsequently, Cu+ reverts to Cu2+ due to the inherent instability of Cu+ and the difference in coordination geometry relative to Cu2+ [66,67].
Once the cycle is completed, it repeats through step 3, step 4, and step 5, with the successive reduction and re-oxidation of copper progressively damaging the crystal structure until it ultimately loses its order and catalytic activity. Among these, step 3 emerges as the critical stage in the context of crystal stability.
In this study, the catalytic reaction mechanism was investigated by selecting a representative fragment of the crystal structure (Figure 13) rather than employing periodic surface models. This approach is justified because the active sites involved in the catalytic cycle are highly localized in the metal coordination environment and its immediate ligands.
Consequently, a molecular fragment extracted from the crystallographic structure is sufficient to capture the essential electronic and steric features governing the catalytic reaction mechanism [68,69]. Moreover, catalytic activity does not arise from delocalized band structures or long-range surface effects but rather from well-defined coordination sites where edge effects are absent [70]. From a kinetic perspective, the IRC mechanism (Figure ) involves two transition states (TS1 and TS2) and an energy valley where the molecule stabilizes in its catalytically active form. The activation energy (Ea) is sufficiently low to allow the system to proceed to the TS2 intermediate, and after electron injection, hydrogen is generated through the formation of hydrides that combine with either H+ or water. The copper center then returns to its original oxidation state; however, this repetitive redox cycle progressively disrupts the crystal structure during steps 3 to 5.
The CuHCF cluster contains five hexacyanoferrate moieties (5 × [Fe(CN)6]6−), one Cu center (Cu2+), and seventeen sodium counterions (17 Na+). Therefore, the overall charge of the cluster is −11, obtained by charge balance as follows: 5 × (−6) = −30 (from [Fe(CN)6]6−), +2 (from Cu2+) gives −28, and +17 (from 17 Na+) gives a net charge of −11. Accordingly, the model can be summarized as Na17Cu[Fe(CN)6]5 with an overall charge of −11.
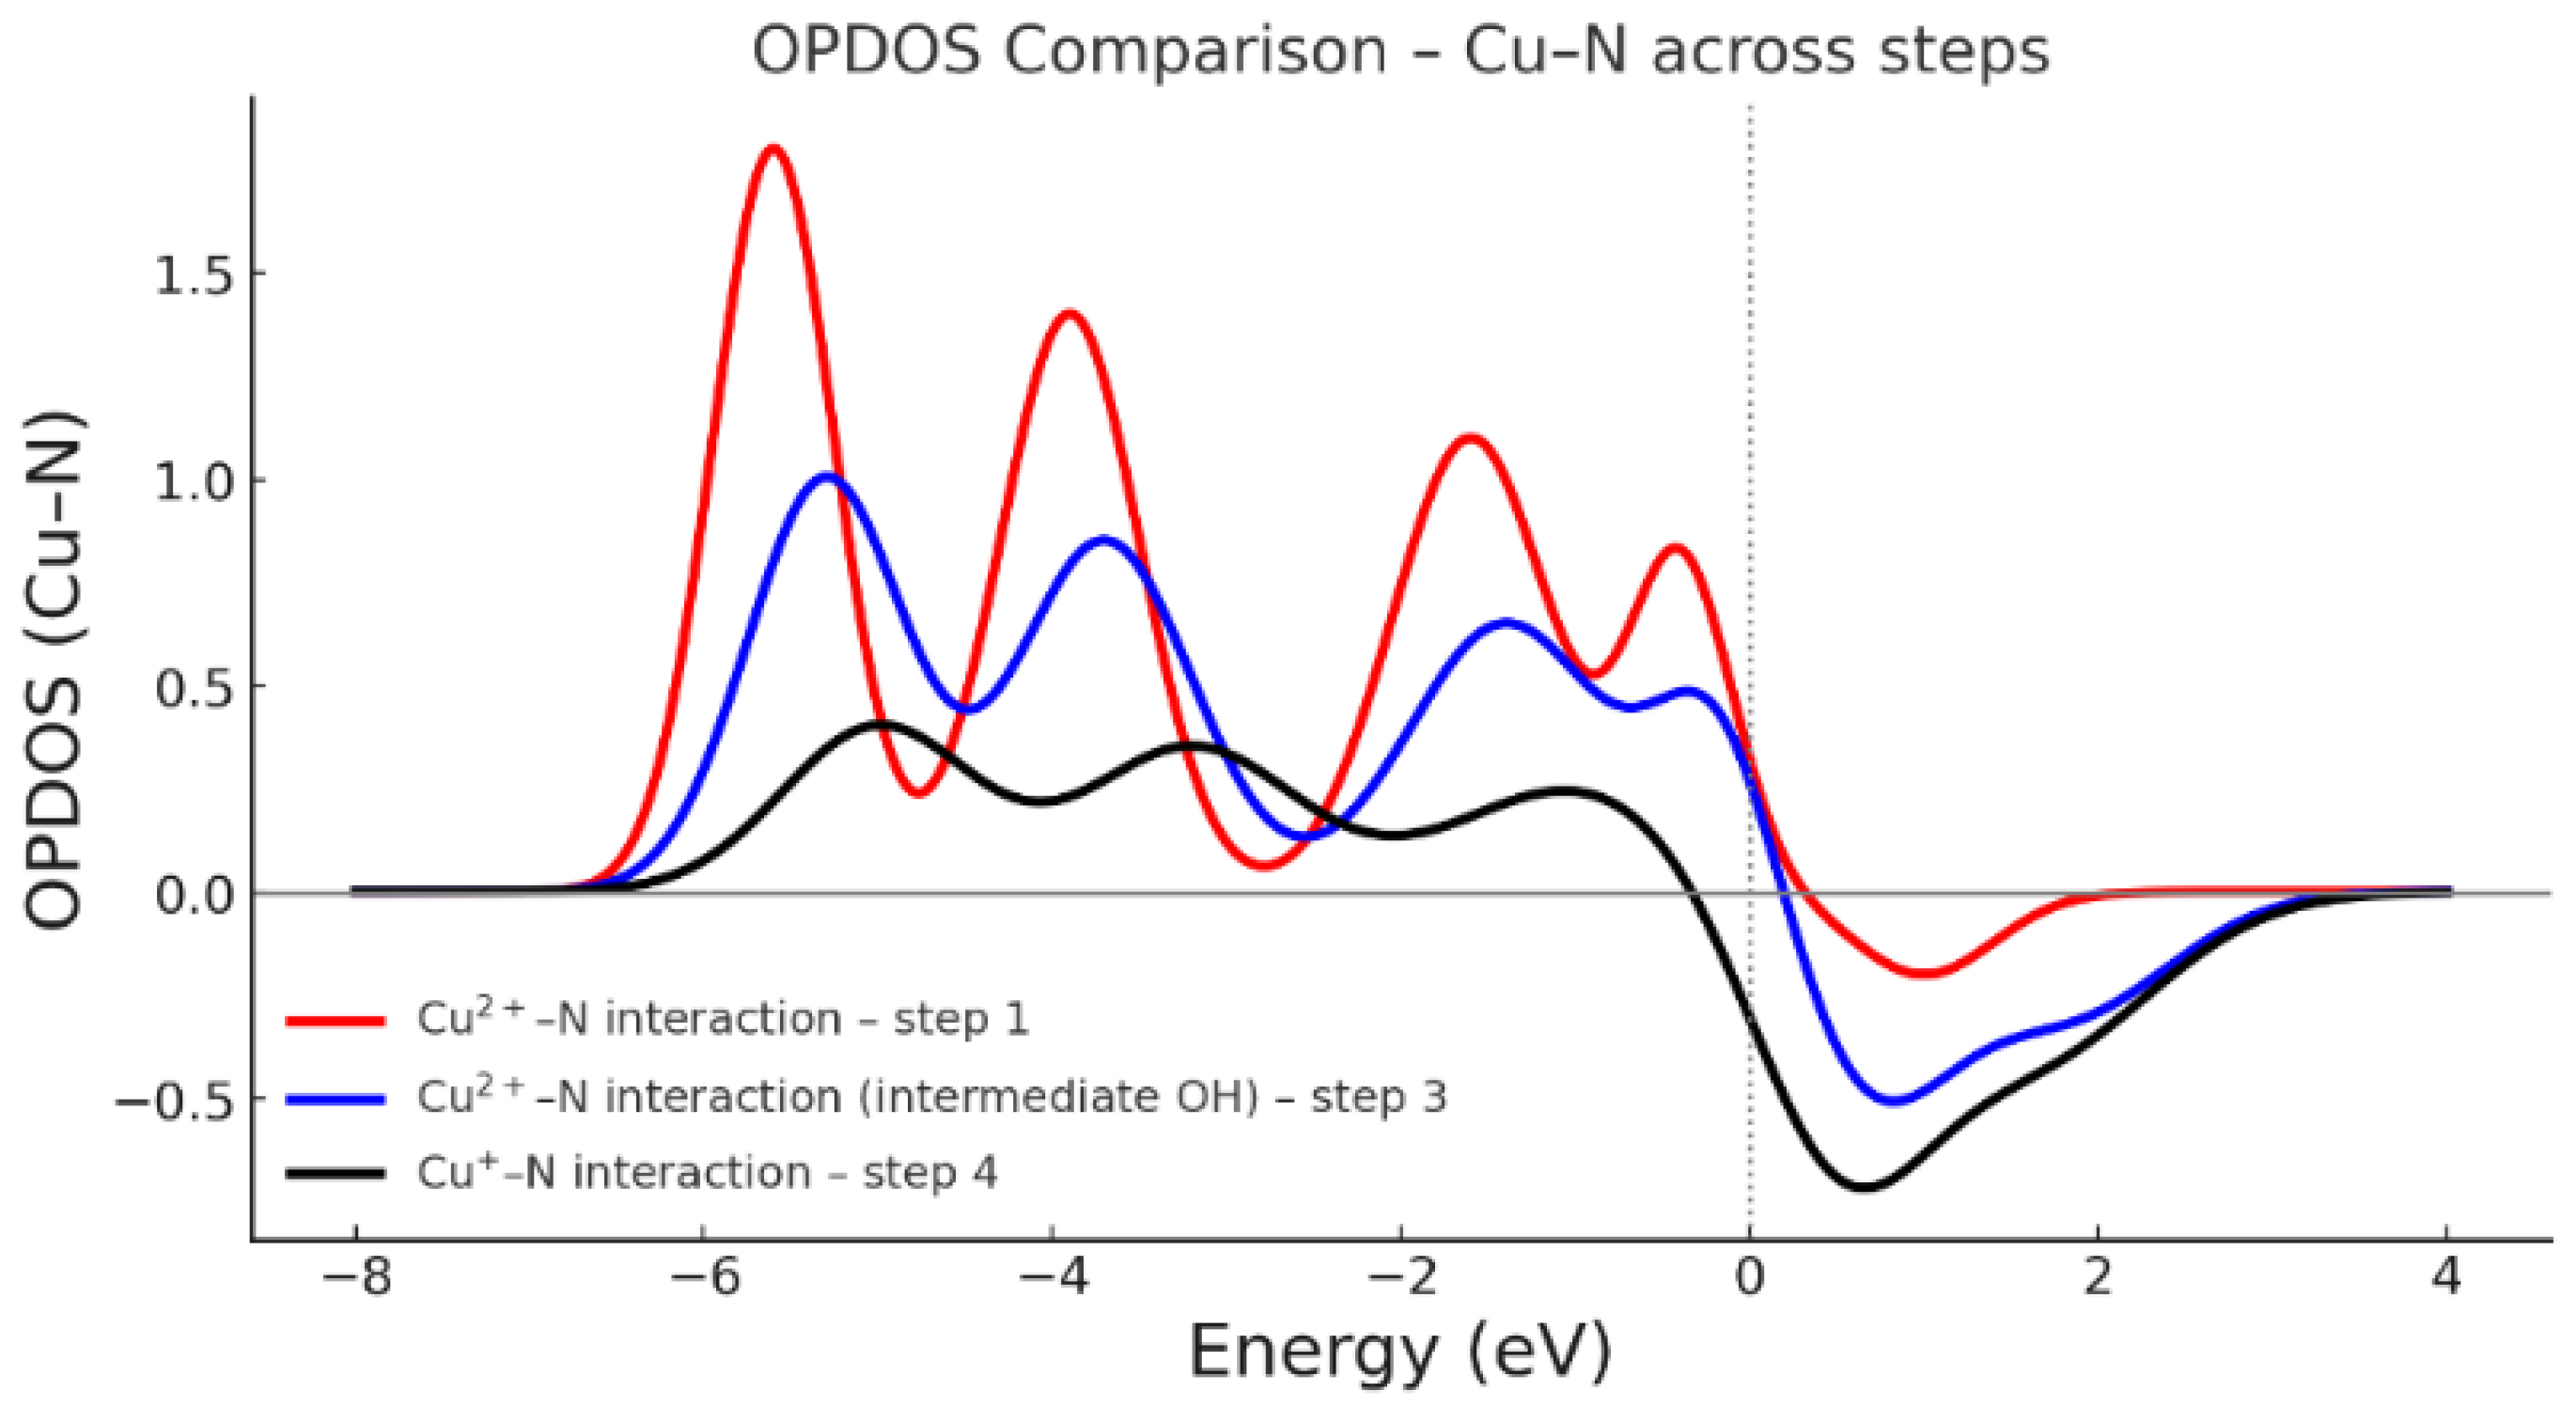
Figure 15 shows the evolution of the electronic interactions between copper and the hexacyanoferrate complex (CuHCF) across three different stages, analyzed through the OPDOS curves. In the red curve, corresponding to the system with copper in the 2+ oxidation state, a strong density of bonding states is observed between approximately −6 eV and −3 eV, reflecting significant overlap between the copper d orbitals and the nitrogen p orbitals of the hexacyanoferrate. The pronounced intensity of these peaks indicates that the Cu2+–N interaction is stabilized by very strong coordination bonds. Furthermore, in this initial stage, no occupied states are detected near the Fermi level, suggesting high electronic stability of the system.
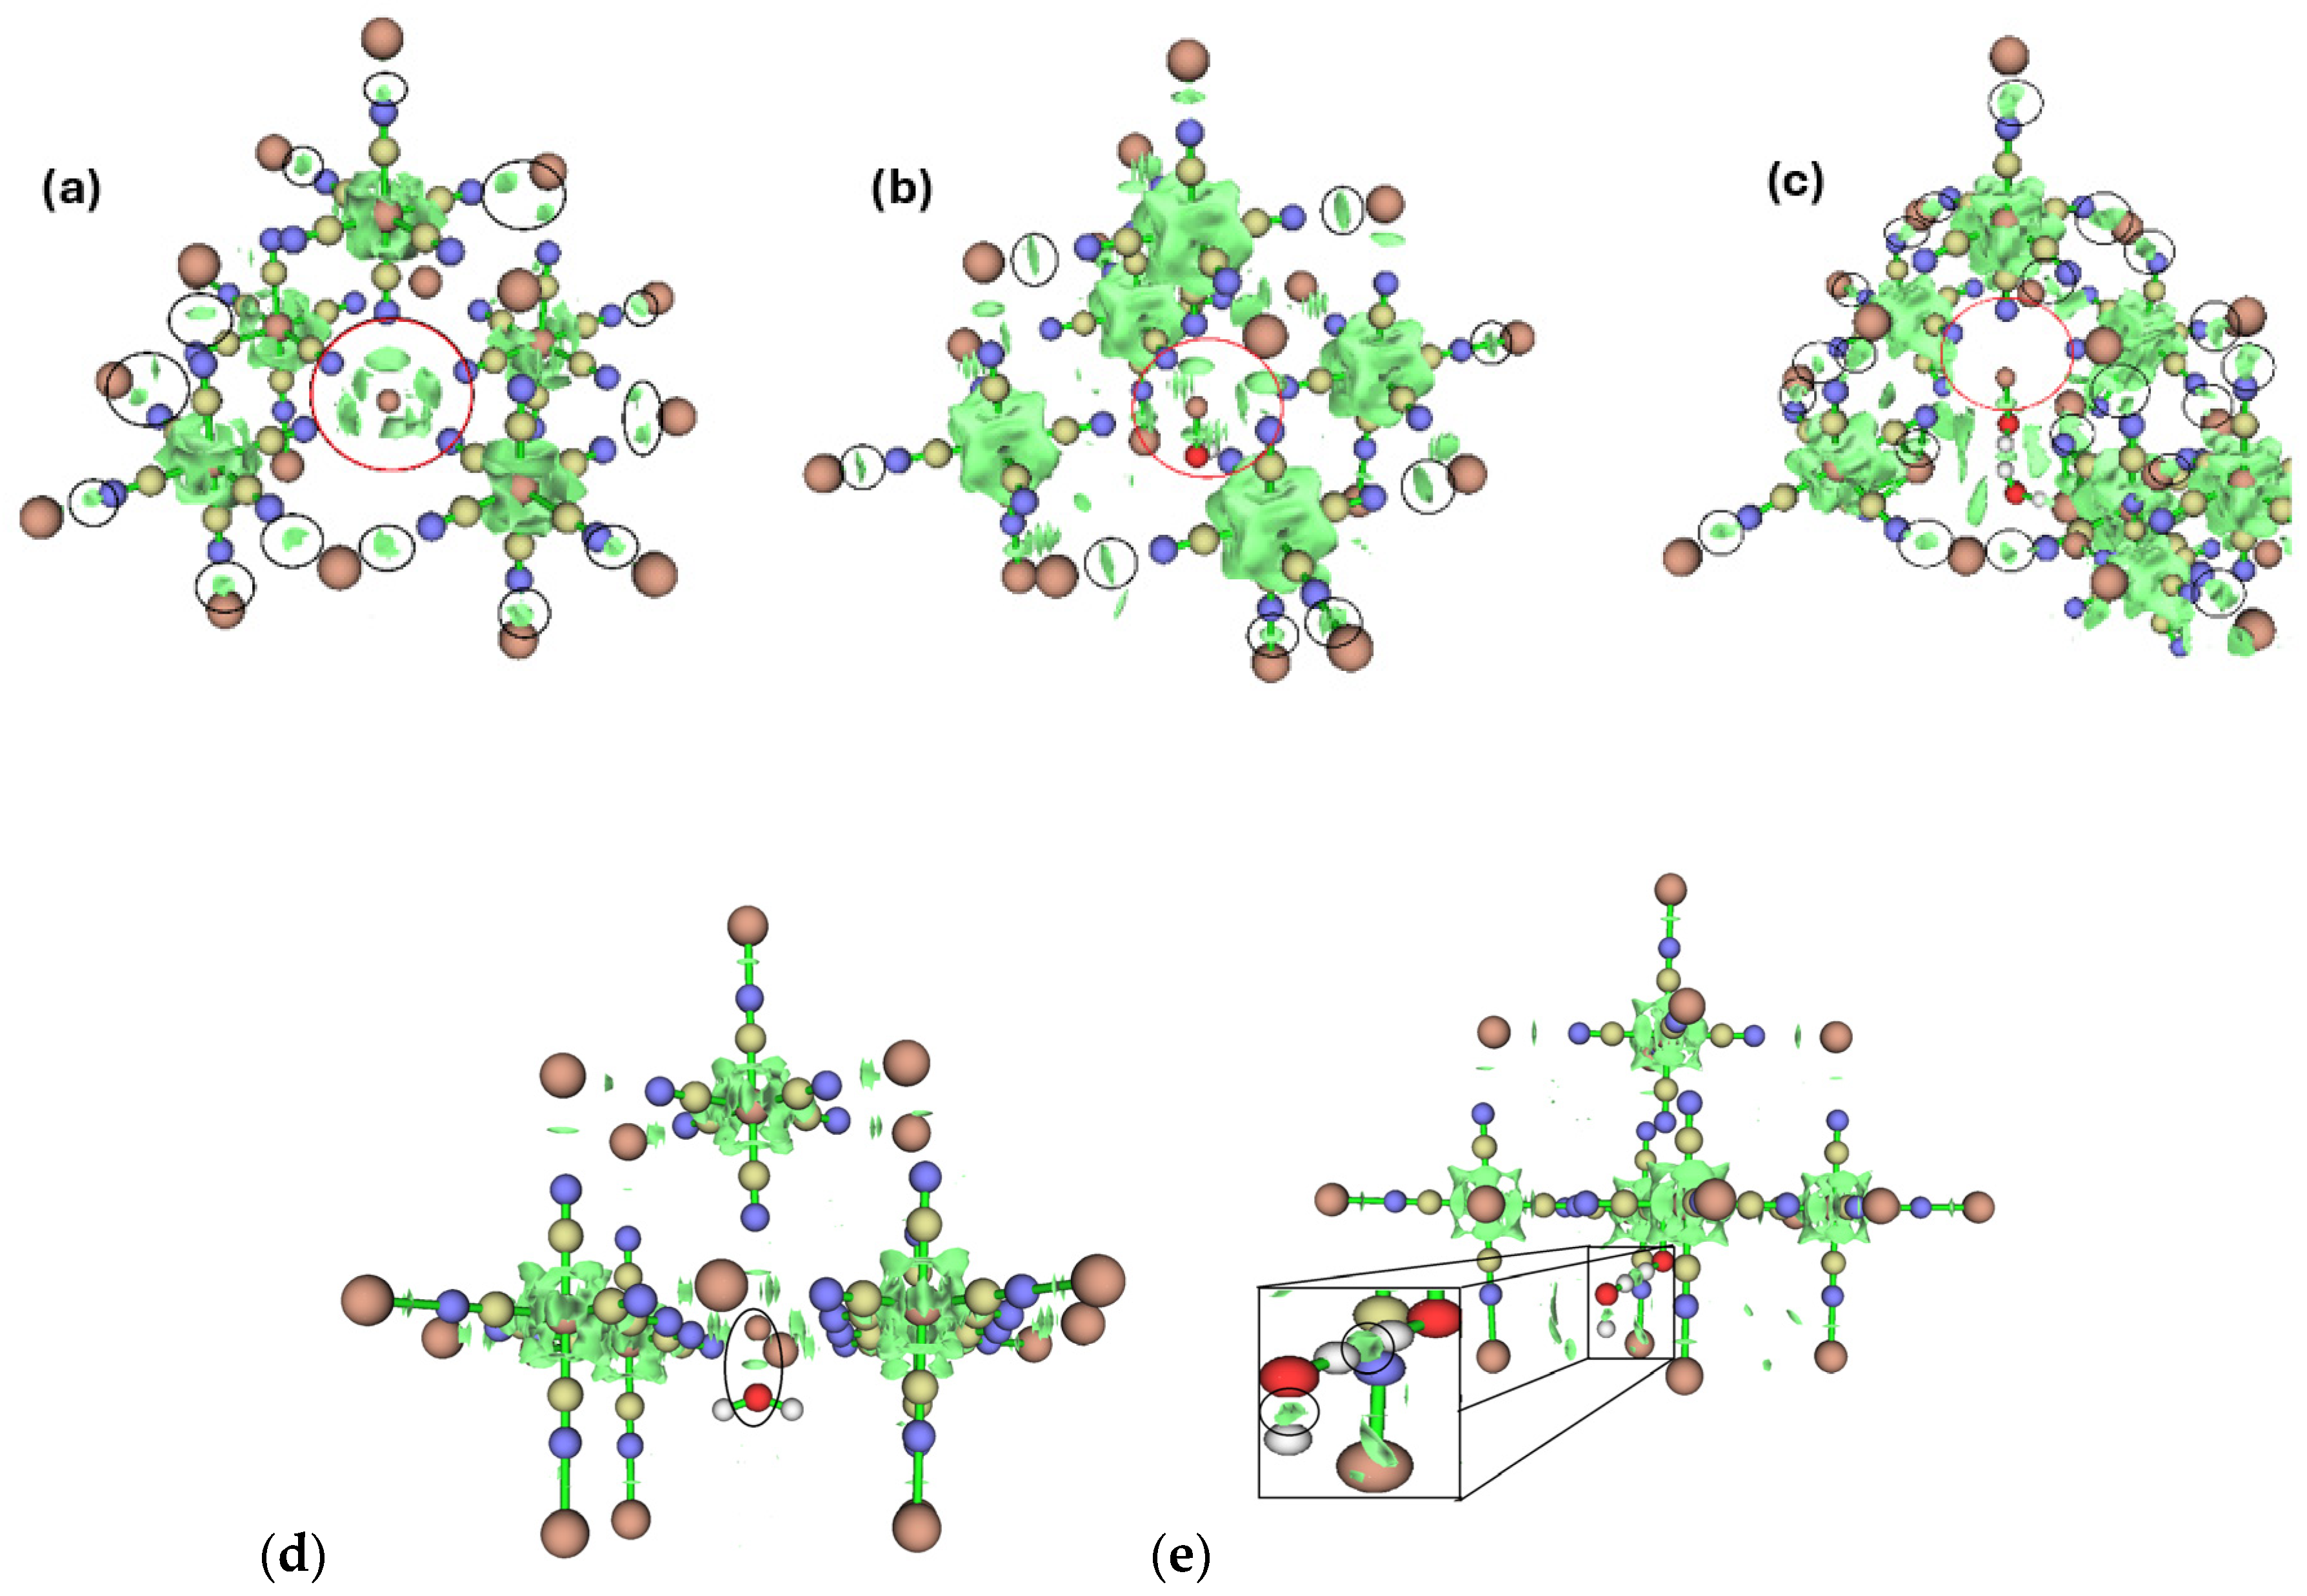
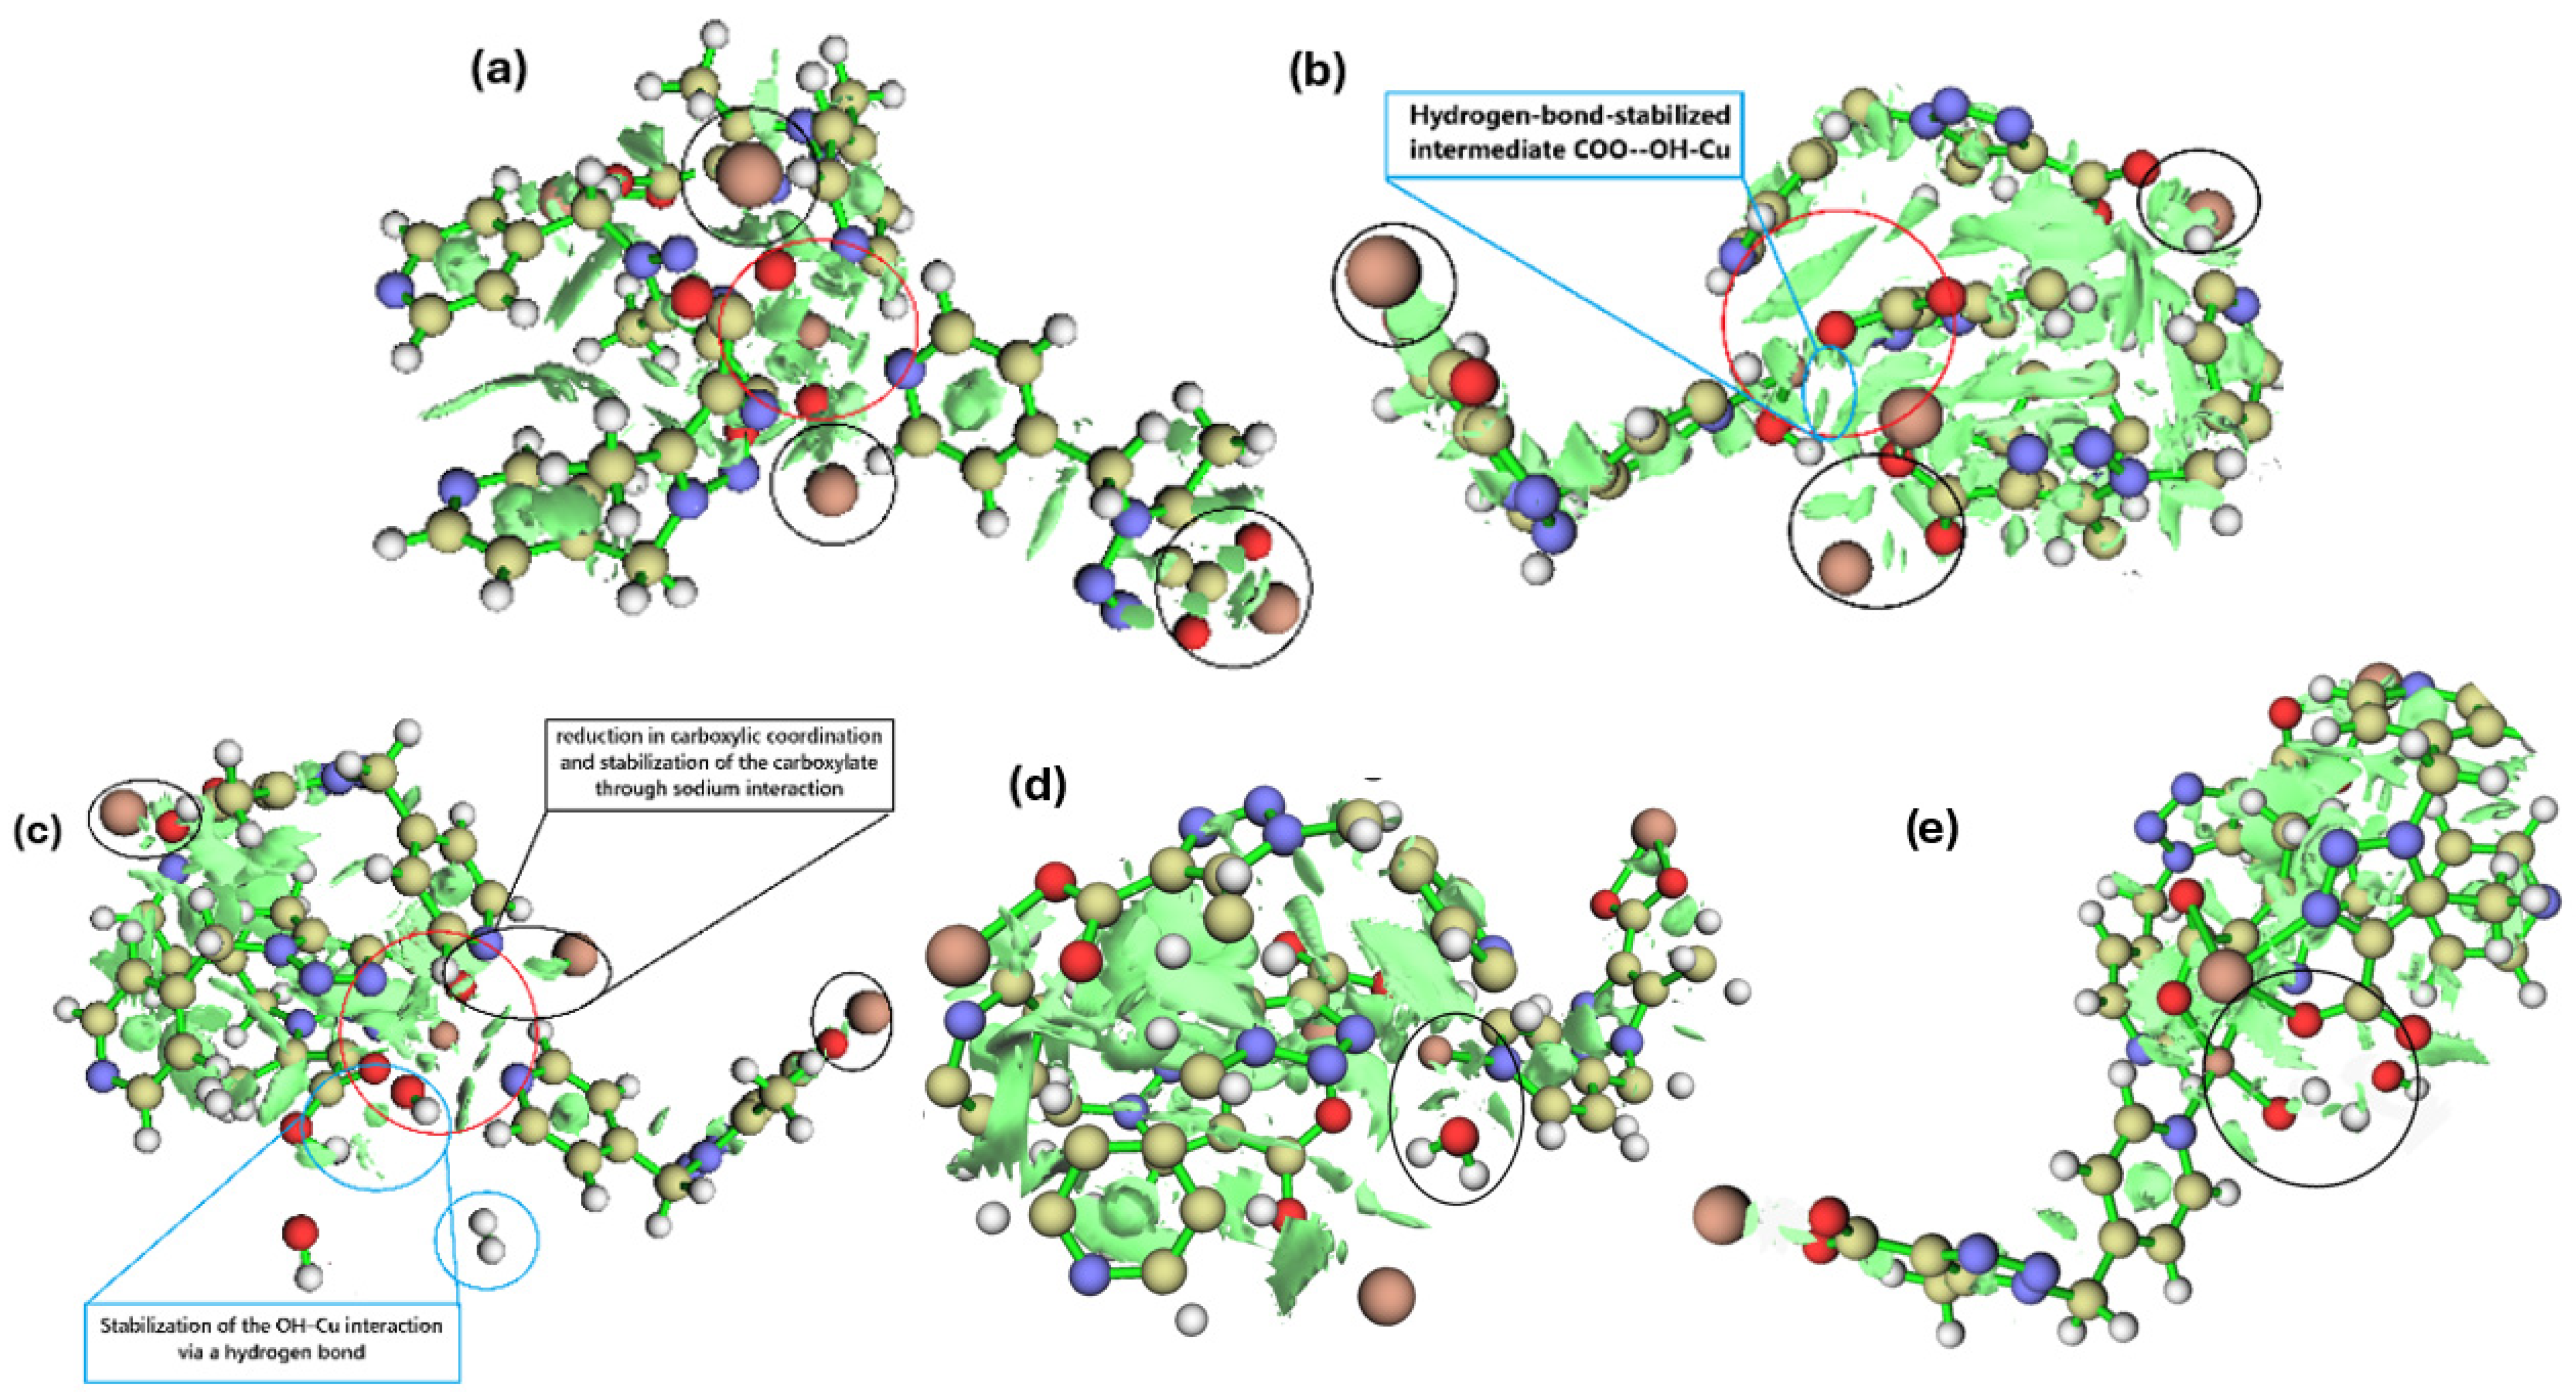
The RDG plots (Figure 16) are consistent with these results, displaying intense green iso-surfaces surrounding the copper atom (Figure 16a), which, together with the OPDOS analysis, confirm the presence of a stable electronic coupling [71]. It is worth noting that, although most of the contributions in this curve are bonding in nature, a slight presence of antibonding states appears in the positive energy region (Figure 15). This indicates that, despite the strong coordination, empty orbitals are available that could participate in electron transfer processes under reactive conditions; once populated, these orbitals may destabilize or weaken the interaction strength due to their antibonding character.
The blue curve in Figure 15 corresponds to the same system with Cu2+ but in its active state, i.e., as an intermediate (with a coordinated OH− molecule) (Figure 12, step 3). This species rapidly converts into Cu+, which is inherently unstable and whose molecular geometry is not compatible with that of Cu2+ [66,67]. Such a reduction acts as the trigger for the catalytic process. In addition, the incorporation of the incoming water molecule alters the electronic distribution around the metal center, as evidenced by a decrease in the intensity of the bonding peaks compared with the red curve. This indicates that water incorporation partially weakens the orbital overlap between copper and nitrogen (Figure 15).
This effect can be attributed to a redistribution of electron density and an increase in antibonding orbital contributions. The RDG iso-surfaces (Figure 16b) show a reduction in the intensity of Cu–N interactions compared with the previous complex, highlighting a weaker coupling strength with the metal. Although the interaction regions are still present, their density is lower, and signs of a more distorted coordination environment appear. Moreover, in this configuration, the antibonding contribution is slightly more pronounced than in the previous case, as indicated by the increased presence of positive states in the OPDOS (Figure 15). This reflects that the presence of water not only reduces bond strength but also facilitates the emergence of antibonding orbitals, thereby imparting catalytic capability to the molecule. Such catalytic activity arises because water introduces slight contributions near the Fermi level (Figure 15), which may promote electron transfer processes in subsequent reactive steps.
Finally, the black curve represents the system at the onset of catalytic activity (Figure 12 and Figure 14), that is, when the complex is reduced to Cu+. The Cu+–N coupling is much weaker compared with the previous configurations. In addition, the appearance of peaks near and slightly above the Fermi level indicates an increase in antibonding orbital contributions, suggesting that the system becomes more susceptible to electron acceptance and therefore exhibits higher electronic reactivity. The transition from Cu2+ to Cu+ is thus associated with a change in the nature of the Cu–N interaction, shifting from a strongly coordinated and stable regime to one where the antibonding contribution is more pronounced.
Linking the RDG planes to the charge analysis provides a coherent picture of how the Cu–ferrocyanide coordination environment evolves during collapse. The NBO atomic charges on the nitrogen donor atoms of the ferrocyanide ligands show a clear progression along the distortion coordinate: in the configuration where the donor N sites are closest to the central Cu atom (i.e., prior to the onset of the process), these N atoms carry charges of approximately −0.6, whereas upon progression of the crystal collapse the charges on the hexacyano groups shift toward substantially less negative values, reaching ~−0.1. This systematic change is consistent with a pronounced reduction in local polarization/charge localization at the donor sites as the Cu–ligand environment reorganizes. Consistently, the corresponding RDG planes show that the interaction region between Cu and the donor site is more clearly defined in the short-distance configuration and becomes progressively weaker as the structure collapses and the Cu···N contact is modified. Importantly, RDG is not interpreted as a direct “charge” metric; rather, it complements the NBO analysis by providing a real-space map of where the interaction is manifested and how it evolves, while the NBO charges quantify the accompanying electronic polarization around the Cu-centered coordination environment.
At this final stage, the RDG plots (Figure 16) confirm the almost complete loss of Cu–N interactions, as the coupling regions observed in earlier stages practically vanish. In contrast, RDG iso-surfaces associated with interactions between the [Fe(CN)6]4− groups and sodium cations increase significantly (Figure 16c), indicating that, upon the loss of Cu–N coordination, the system tends to stabilize through new electrostatic interactions with Na+ to compensate for the structural instability caused by copper reduction.
Although the redox transition from Cu2+ to Cu+, which enables electron transfer for hydrogen formation (Figure 12 and Figure 14, steps 4 and 5), is fast and reversible, each cycle progressively weakens the structure. The material is still able to sustain hydrogen production until a critical point is reached where the CuHCF lattice partially collapses. This collapse does not imply an immediate breakdown of the entire framework but begins with local deformations in the CuN6 coordination environment (Figure 16c). During reduction, the [CuN6] octahedra undergo significant tetragonal distortions associated with Jahn–Teller effects [72,73] where the ideal octahedral symmetry is broken and some Cu–N bonds elongate while others contract. As the system cycles between Cu2+ and Cu+ states, these distortions gradually accumulate, compromising the continuity of the crystalline network and generating internal strain that weakens connectivity between the metal centers and the ligands. When the system returns to the Cu2+ state after several cycles, the original geometric structure is not fully recovered, and the unit cell adopts an unstable, strained configuration with local defects and accumulated distortions (Figure 16c).
At this advanced stage, the ferrocyanide groups begin to strengthen their interactions with sodium cations in the environment as a means of stabilizing the lattice in response to the loss of structural symmetry. This phenomenon is evidenced by the RDG plots (Figure 16c), which reveal additional attraction regions associated with the coordination between [Fe(CN)6]4− groups and Na+ ions. Such interactions act as a compensation mechanism but come at the expense of introducing further strain and distortions into the crystal lattice, which, upon accumulating over repeated hydrogen evolution cycles, ultimately lead to the partial and irreversible collapse of the CuHCF framework.
The proposed mechanism involves not only the CuHCF compound but is also applicable to the CCCu complex (Figure 17, Figure 18 and Figure 19). The mechanism is consistent with the experimental results and explains why CuHCF decomposes rapidly, whereas CCCu retains catalytic activity with greater resistance to degradation (Figure 19).
Experimental data show that the exchange current density for HER confirms the higher intrinsic activity of CCCu (|i0, H2| = 0.062 A/m2) compared with CuHCF (0.019 A/m2), and stands in sharp contrast to titanium, whose HER activity in brine is practically negligible (0.001–0.01 A/m2) [48].
The Tafel slope observed for CuHCF, close to (|bH2| ≈ 222 mV/dec), can be interpreted in terms of the reaction mechanism involved in hydrogen evolution. This high value is generally associated with a Volmer–Heyrovsky mechanism, in which the Volmer acts as the rate-determining step of the overall process [74]. In this scenario, the kinetics are controlled by the difficulty of the material in facilitating the initial charge transfer and the formation of the adsorbed intermediate, which is reflected in the steep slope. A more precise explanation of this behaviour is essential, as it allows the correlation of kinetic parameters with the nature of the active sites of CuHCF and its ability to mediate the cleavage of the O–H bond in saline media.
The comparative stability between both compounds is reflected in the experimental values of the oxidation/dissolution exchange currents: i0, M(CuHCF) = 0.740 A/m2 versus i0, M(CCCu) = 0.395 A/m2. The higher i0, M of CuHCF indicates a greater tendency to oxidize/dissolve within the explored potential range, consistent with the proposed mechanism involving Cu2+ → Cu+ reduction coupled to the progressive rupture of Cu–N/CN bonds. In contrast, CCCu retains its cathodic response due to its stronger and more flexible crystalline configuration (absence of labile CN-type ligands), the lower dissolution rate inferred from i0, M, and the more sterically hindered access to the metal center imposed by the ligand orientation (Figure 19).
Both materials exhibit relatively good corrosion potentials (−23 to −40 mVSHE) compared with titanium under neutral conditions (−456 mVSHE in 0.6 M NaCl) [35], although CuHCF remains intrinsically unstable due to the presence of antibonding states. Once again, the experimental data agrees with the proposed theoretical mechanism, in which CuHCF is active for HER.
Regarding electronic conduction, in CuHCF and CCCu, it is mainly governed by the presence of polarons, which are closely linked to its structural fragility. In this compound, charge is transported predominantly through polaron hopping [75] between redox centers via C≡N–Cu/Fe bridges. This mechanism implies that electronic transport is localized rather than delocalized, making electron mobility highly dependent on the integrity of the bonding network. As mentioned earlier, the OPDOS analysis reveals the presence of antibonding states (σ* Cu–N/π* C≡N) close to the Fermi level, which become populated during the electrocatalytic process (steps 3–5) (Figure 14 and Figure 15). The occupation of these states progressively weakens the Cu–N/C≡N bonds that provide rigidity to the crystal, leading to the breaking of CN connectivity. Consequently, σ bonds collapse, charge transfer pathways are blocked, and/or charge-transfer resistance increases, ultimately resulting in a larger Tafel slope.
The proposed mechanism applies equally to both CuHCF and the CCCu complex, although kinetic differences can be observed in the IRC (Figure 17). The potential well in CCCu is much more stable than in CuHCF, a feature attributable to the structural composition of its ligands, as illustrated in Figure 17, Figure 18, Figure 19 and Figure 20, respectively. The incorporation of a water molecule into the crystalline lattice is stabilized by the non-coordinating carboxylate oxygen, making the intermediate of this compound (Figure 17, step 3; Figure 19) more stable. As a result, the Cu2+ → Cu+ transition requires a somewhat higher second activation energy compared to CuHCF, which partly contributes to reducing the instability of the complex during the redox process.
Similar to CuHCF, steps 1 and 2 (Figure 12 and Figure 17) are non-catalytic and correspond to the formation of the catalytic species in step 3, which subsequently produces hydrogen through the Cu2+/Cu+ redox transition. The catalytic cycle then repeats between steps 3 and 5 for both compounds (Figure 12 and Figure 17). Accordingly, the proposed mechanism, consistent with the experimental data, suggests an unusual catalytic behavior that does not fit within current definitions of electrocatalysts. For this reason, we therefore define these materials as “semi-electrocatalysts”: The term semi-electrocatalyst is proposed to describe materials that exhibit partial and/or transient catalytic activity driven by controlled structural degradation during the electrochemical process. Unlike conventional catalysts, whose chemical structure remains stable throughout electrolysis, semi-electrocatalysts undergo an initial activation followed by a progressive modification of their coordination network, enabling them to maintain both a moderate catalytic function and an anticorrosive role in aggressive media such as seawater (see Figure 18). This novel conceptual proposal emphasizes that, in their initial state, such materials exhibit no catalytic activity, yet upon activation, they temporarily function as catalysts before their structural framework collapses.
In the same way as in the hydrogen formation mechanism study with CuHCF, a representative unit of the CCCu crystal was selected for modelling (Figure 18). This approach allowed us to verify whether both compounds follow the same reaction mechanism and to clarify the structural features responsible for their differences. Figure 19 presents the Intrinsic Reaction Coordinate (IRC) for hydrogen production in CCCu, showing the energy profile along the reaction pathway. The curve highlights the formation of a hydrogen-bond-stabilized intermediate, followed by the evolution of H2. This analysis confirms that, although CCCu and CuHCF share similar mechanistic steps, their energetic barriers and stabilization effects differ, reflecting the influence of coordination environment and crystal structure on catalytic activity.
For the CCCu cluster model, the system includes four sodium ions (4 Na+, total +4) coordinated to four deprotonated carboxylic acid groups (4 × COO−, total −4), accounting for internal charge compensation within the ligand environment. In addition, the cluster contains a central copper ion (Cu2+). Therefore, the overall charge of the CCCu cluster is +2 (net 0 from the 4 Na+/4 COO− pairs, plus +2 from Cu2+).
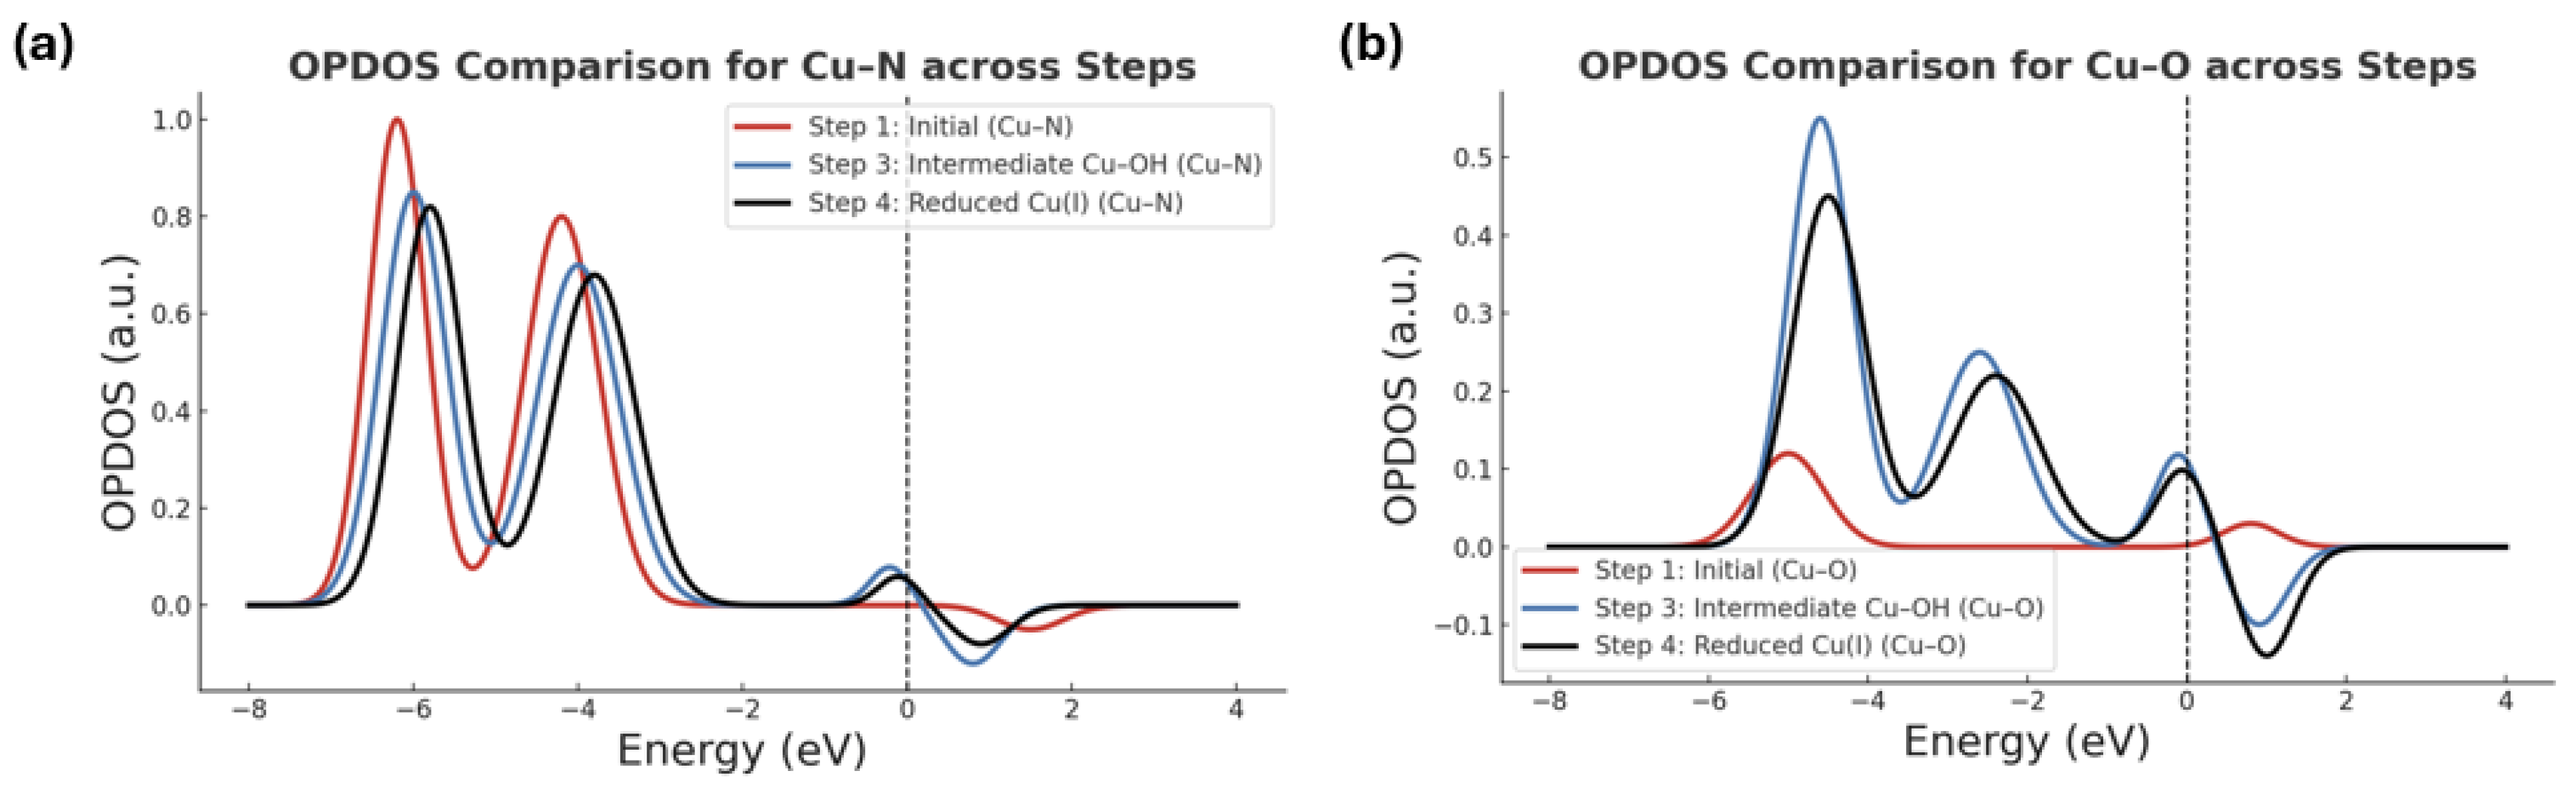
To gain further insights into the electronic structure of CCCu during the hydrogen evolution pathway, an OPDOS analysis was performed (Figure 20). The comparison of Cu–N and Cu–O interactions across the different reaction steps provides a detailed picture of how ligand coordination governs stability and reactivity. The strong Cu–N overlap in the initial stage explains the preserved geometry, while the progressive involvement of Cu–O interactions in the intermediate and reduced states reflects the electronic redistribution accompanying hydrogen evolution.
The OPDOS results (Figure 20 obtained for the CCCu complex reveal an electronic distribution consistent with greater stability in the initial stage (Step 1; Figure 17 and Figure 19). The OPDOS for Cu–N interactions (Figure 20a) display intense bonding peaks between −6 eV and −3.5 eV, associated with strong Cu(d)–N(p) orbital overlap originating from the pyridine ligands, while the Cu–O contributions are comparatively weaker (Figure 20b) and shifted to slightly lower energies. This indicates that the coordinating carboxylate groups contribute to the stabilization of the electronic environment but play a secondary role relative to the pyridine units. Such a configuration accounts for the excellent coupling between copper and the N ligands, preserving a stable initial geometry.
As the system progresses to the intermediate stage (Step 3; Figure 17 and Figure 19), corresponding to water incorporation and the formation of the Cu–OH intermediate, the OPDOS (Figure 20) reveal a subtle redistribution of electronic density, with a slight decrease in the intensity of the Cu–N peaks and the appearance of states near the Fermi level, primarily associated with Cu–O contributions. This indicates that pyridine coordination remains virtually unaltered, thereby stabilizing the intermediate, while the coordinating carboxylate oxygens undergo only minor displacements due to partial interaction with Na+ cations. To visualise the structural changes associated with hydrogen evolution in the CCCu complex, electron density difference maps were generated for the key reaction steps (Figure 21). These maps highlight the redistribution of charge density around the Cu–N and Cu–O coordination spheres. The RDG analysis (Figure 21b) supports this trend, showing well-defined iso-surfaces with no significant loss of symmetry.
Experimentally, this stability is reflected in a low Tafel slope (≈148 mV/dec), indicating an efficient and favourable charge-transfer mechanism, together with relatively high exchange current densities (≈0.062 A/m2). These features are consistent with the presence of accessible electronic states and a more durable structural framework. They undergo slight modifications while maintaining their bonding character, whereas the Cu–O contributions show a modest increase in antibonding density near the Fermi level. This behavior reveals that the electronic reorganization is mainly concentrated within the coordination sphere of the carboxylate oxygens, while the rigid pyridine ligands contribute significantly to the overall structural stability. This interpretation is consistent with the thermal stability observed in the TGA analysis, where the complex retains its structural integrity up to approximately 350 °C, as well as with the low dissolution current measured experimentally (i0, M ≈ 0.395 A/m2) and the reduced degradation observed after multiple operating cycles.
However, despite this high initial stability, the CCCu complex is intrinsically unstable and will eventually collapse over time. The progressive filling of Cu–O antibonding states, as evidenced in the OPDOS near the Fermi level (Figure 20b), gradually weakens the lattice rigidity and reduces the strength of the Cu–O bonds after numerous redox cycles. In addition, partial intercalation of Na+ ions into the coordination environment introduces small cumulative distortions that, although minimal in each cycle, generate increasing local strain. This is further aggravated by the electronic and mechanical fatigue associated with the continuous Cu2+ ↔ Cu+ transitions, which induce repetitive elongation and contraction of the bonds, particularly at the carboxylate oxygens, progressively compromising structural integrity. These perturbations, though slow, are unavoidable and accumulate until they generate microfractures within the lattice, loss of electronic connectivity, and ultimately the collapse of the crystalline network.
Taken together, the OPDOS data (Figure 20), RDG analysis (Figure 21), IRC results (Figure 19), and experimental evidence confirm that CCCu exhibits an optimized electronic design, in which the dominant contribution of Cu–N interactions stabilizes the Fermi level and preserves structural integrity, while Cu–O interactions, although secondary, govern local flexibility and modulate the dynamics of degradation. Nonetheless, long-term collapse remains inevitable due to the cumulative effects of antibonding state filling. Therefore, CCCu can be considered a semi-electrocatalyst according to our definition: it requires activation by the medium, maintains activity and stability for part of its lifetime, but undergoes unavoidable degradation over time. This positions both materials as representatives of a new type of catalyst, potentially applicable in scenarios where hydrogen production is required only for a defined period.

4. Conclusions

This study establishes copper-based coordination compounds, specifically a triazole-derived coordination polymer (CCCu) and a Prussian Blue analogue (CuHCF), as pioneering materials for hydrogen production in saline environments. Structural and spectroscopic analyses (FTIR, TG/DSC, XRD, SEM/EDS) confirmed successful coordination of organic ligands to copper centres, thermal stability up to ~120 °C for CCCu and ~150 °C for CuHCF, and distinct morphologies: a laminar, high-surface-area structure in CCCu and a mesoporous, nanostructured spherical architecture in CuHCF. These features directly influenced their electrochemical behaviour.
Electrochemical testing in 0.5 M NaCl revealed that CuHCF exhibited superior HER kinetics, with higher exchange current densities (0.019 A/m2) and lower Tafel slopes (~220 mV/dec) compared to CCCu and commercial titanium (Ti Gr.2). In contrast, CCCu showed lower intrinsic catalytic activity but greater corrosion resistance and structural retention under prolonged operation. When compared to conventional steels (A36, AISI 1020, A630-420H), which displayed rapid dissolution (icorr 1.36–3.02 A/m2, Ecorr −221 to −320 mVSHE), both CCCu and CuHCF offered a unique compromise of catalytic activity and durability, bridging the gap between catalytically active yet unstable steels and catalytically inert but stable titanium.
A superposition model based on mixed-potential theory enabled deconvolution of partial current contributions for HER, ORR, and CCOR, providing a rigorous kinetic framework to assess performance in complex saline systems. Importantly, DFT calculations provided mechanistic insight into the role of metal–ligand interactions and electronic redistribution in governing catalytic activity and stability. The computed Gibbs free energy of hydrogen adsorption supported the experimentally observed differences between CCCu and CuHCF, linking structural features to catalytic behaviour. These results demonstrate how the integration of computational and experimental approaches enables a coherent framework for understanding activity–stability trade-offs in coordination-based electrocatalysts.
The central conceptual advance of this work is the definition of “semi-electrocatalysts”: materials that initially require activation, generate transient catalytic species capable of sustaining repeated catalytic cycles, but ultimately undergo structural degradation that limits their lifetime. Within this framework, CCCu and CuHCF emerge as the first representatives of this new class.
Despite their moderate catalytic activity and inherent structural instability, the copper-based coordination compounds investigated here can be realistically envisioned for short-term or niche applications where cost, abundance of precursors, and system simplicity are prioritized over long-term durability. In this context, their role as semi-electrocatalysts positions them as suitable candidates for intermittent hydrogen generation at low current densities, particularly when integrated with solar-driven systems in coastal regions. Such applications would benefit from direct utilisation of saline water without extensive pre-treatment, thereby reducing operational costs. While these materials are not competitive with state-of-the-art electrocatalysts for continuous, large-scale hydrogen production, they open a pathway towards decentralised, low-cost solutions where controlled degradation can be tolerated under defined operational lifetimes.
Finally, the comparative evaluation underscores a key trade-off: CCCu provides higher structural stability and corrosion resistance, while CuHCF delivers faster HER kinetics and superior mass transport properties. Together, they demonstrate that copper-based coordination compounds can serve as rational precursors for the design of next-generation semi-electrocatalysts, paving the way for cost-effective, solar-driven seawater electrolysis technologies where durability, selectivity, and catalytic efficiency must be carefully balanced.

Supplementary Materials

The following supporting information can be downloaded at: https://www.mdpi.com/article/10.3390/pr14030423/s1, Table S1: Negative frequencies of transition states.

Author Contributions

Conceptualization, F.M.G.M., A.S., P.Z., M.B., D.G., N.T., I.B. and V.M.J.-A.; methodology, F.M.G.M., P.Z., M.B., I.B. and V.M.J.-A.; software, P.Z., F.M.G.M., L.C., A.S. and M.B.; validation, F.M.G.M., N.T., D.G. and V.M.J.-A.; formal analysis, D.G., N.T., P.Z. and F.M.G.M.; investigation, F.M.G.M., I.B., L.C., M.B., V.M.J.-A., A.S. and P.Z.; resources, F.M.G.M.; data curation, A.S. and F.M.G.M.; writing—original draft preparation, F.M.G.M., P.Z., N.T., M.B., L.C., A.S., D.G., I.B. and V.M.J.-A.; writing—review and editing, F.M.G.M., P.Z., M.B., I.B. and L.C.; visualization, F.M.G.M.; supervision, F.M.G.M.; project administration, F.M.G.M. All authors have read and agreed to the published version of the manuscript.

Funding

This research received no external funding.

Data Availability Statement

The original contributions presented in this study are included in the article/Supplementary Materials. Further inquiries can be directed to the corresponding author.

Acknowledgments

The authors would like to thank the support of ANID-Chile through the research project FONDECYT Iniciación 11230550; and this work was supported by ANID–Chile, through the Solar Energy Research Center (SERC Chile), under the National Research Centers of National Interest Program, grant CIN250043. Additionally, V. Jimenez-Arevalo appreciates the financial assistance provided by the Vicerrectoria de Postgrado from the Universidad de Santiago de Chile and postdoctorate project DICYT N° 022442PC, Vicerrectoría de Investigación, Innovación y Creacion. This work further benefited from the equipment acquired through ANID’s Concurso Equipamiento Científico y Tecnológico FONDEQUIP Mediano 2022, EQM 220028 “Adquisición de un Microscopio Electrónico FE-SEM para el fortalecimiento de la investigación, vinculación y docencia de pre y postgrado de la Universidad de Antofagasta. Thank MINEDUC-UA proyect, Code ANT 22991. We gratefully acknowledge the Laboratory for Electrochemistry, Corrosion and Solar Hydrogen at the Centro de Desarrollo Energético de Antofagasta (CDEA) for providing the infrastructure and technical facilities essential for this study. We also thank the Solar Energy PhD Program at the University of Antofagasta for its academic support throughout the research process.

Conflicts of Interest

The authors declare no conflicts of interest.

References

  1. Shiva Kumar, S.; Himabindu, V. Hydrogen production by PEM water electrolysis—A review. Mater. Sci. Energy Technol. 2019, 2, 442–454. [Google Scholar] [CrossRef] [Scilit]
  2. Yan, Y.; Xia, B.Y.; Zhao, B.; Wang, X. A review on noble-metal-free bifunctional heterogeneous catalysts for overall electrochemical water splitting. J. Mater. Chem. A 2016, 4, 17587–17603. [Google Scholar] [CrossRef] [Scilit]
  3. Jimenez-Arevalo, V.M.; Martin, P.; Pio, E.; Pavez, J.; Aguilar, C.C.; Suryanarayana, C.; Páez, M. Energy Transfer by Mechanical Alloying and Electrocatalytic Performance of the As-Sintered Self-Supported High-Entropy Alloy FeNiCoCuMo. Adv. Eng. Mater. 2025, 27, 2400807. [Google Scholar] [CrossRef] [Scilit]
  4. Swathi, S.; Yuvakkumar, R.; Kungumadevi, L.; Ravi, G.; Velauthapillai, D. Novel synthesis of CuHCF/B-rGO composites for oxygen evolution reaction activity. Sci. Rep. 2024, 14, 25760. [Google Scholar] [CrossRef] [Scilit]
  5. Kim, J.; Seo, J.H.; Lee, J.K.; Oh, M.H.; Jang, H.W. Challenges and strategies in catalysts design towards efficient and durable alkaline seawater electrolysis for green hydrogen production. Energy Mater. 2025, 5, 500076. [Google Scholar] [CrossRef] [Scilit]
  6. Ghosh, A.; TK, S.F.; Ganguly, D.; Sundara, R. CoFePBA Nanosheets on Carbon Nanotubes Coupled with Nickel-Encapsulated Carbon Tubules for Efficient and Highly Stable Overall Seawater Electrolysis. ACS Appl. Energy Mater. 2023, 6, 6080–6090. [Google Scholar] [CrossRef] [Scilit]
  7. Song, X.; Song, S.; Wang, D.; Zhang, H. Prussian Blue Analogs and Their Derived Nanomaterials for Electrochemical Energy Storage and Electrocatalysis. Small Methods 2021, 5, e2001000. [Google Scholar] [CrossRef] [Scilit]
  8. Ibadat, S.; Ashfaq, H.T.; Ruqia; Asghar, M.A.; Haider, A.; Ali, S. Surfactant-Promoted Prussian Blue Analogues Fabricated Electrodes for Electrocatalytic Water Oxidation. Proc. Pak. Acad. Sci. Part B 2023, 60, 549–555. [Google Scholar] [CrossRef] [Scilit]
  9. Sunday, N.F. Emerging Trends in Coordination Polymers and Metal-Organic Frameworks: Perspectives, Synthesis, Properties and Applications. Arch. Org. Inorg. Chem. Sci. 2018, 1, 1–12. [Google Scholar] [CrossRef] [Scilit]
  10. Pan, Y.; Gao, J. Exploring the hydrogen storage capacity, dehydrogenated mechanism, electronic and optical properties of AMMgH3 hydrides for hydrogen storage. J. Energy Storage 2025, 124, 116869. [Google Scholar] [CrossRef] [Scilit]
  11. Yang, F.; Pan, Y. An emerging 2D borophene nanomaterial: Synthesis, properties and hydrogen energy applications. Coord. Chem. Rev. 2025, 545, 217015. [Google Scholar] [CrossRef] [Scilit]
  12. Gao, J.; Pan, Y.; Jain, I.P.; Yang, F.; Zhu, J. Exploring the structure, hydrogen storage capacity and hydrogen storage mechanism of X3N3H6 hydrides for hydrogen storage. Appl. Mater. Today 2025, 44, 102755. [Google Scholar] [CrossRef] [Scilit]
  13. Karmakar, A.; Govindaraja, T.; Baasanjav, E.; Bandyopadhyay, P.; Jin, B.; Sei Park, Y.; Lim, D.-H.; Mun, S. Surface hydroxyl group-enriched nickel cobalt molybdate hydrate for improved oxygen evolution activity in an anion exchange membrane water electrolyzer. Appl. Catal. B 2023, 328, 122504. [Google Scholar] [CrossRef] [Scilit]
  14. Givaja, G.; Amo-Ochoa, P.; Gómez-García, C.J.; Zamora, F. Electrical conductive coordination polymers. Chem. Soc. Rev. 2012, 41, 115–147. [Google Scholar] [CrossRef] [Scilit]
  15. Jiang, W.; Wang, T.; Chen, H.; Suo, X.; Liang, J.; Zhu, W.; Li, H.; Dai, S. Room temperature synthesis of high-entropy Prussian blue analogues. Nano Energy 2021, 79, 105464. [Google Scholar] [CrossRef] [Scilit]
  16. Huang, Y.; Xie, M.; Wang, Z.; Jiang, Y.; Yao, Y.; Li, S.; Li, Z.; Li, L.; Wu, F.; Chen, R. A Chemical Precipitation Method Preparing Hollow–Core–Shell Heterostructures Based on the Prussian Blue Analogs as Cathode for Sodium-Ion Batteries. Small 2018, 14, e1801246. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  17. Desiraju, G.R. Crystal engineering: From molecule to crystal. J. Am. Chem. Soc. 2013, 135, 9952–9967. [Google Scholar] [CrossRef] [Scilit]
  18. Bentz, K.C.; Cohen, S.M. Supramolekulare Metallopolymere: Von linearen Materialien zu infiniten Netzwerken. Angew. Chem. 2018, 130, 15208–15218. [Google Scholar] [CrossRef] [Scilit]
  19. Zhou, H.C.J.; Kitagawa, S. Metal-Organic Frameworks (MOFs). Chem. Soc. Rev. 2014, 43, 5415–5418. [Google Scholar] [CrossRef] [Scilit]
  20. Bergendahl, M.; Narea, P.; Llanos, J.; Pulido, R.; Naveas, N.; Amo-Ochoa, P.; Zamora, F.; Delgado, G.E.; Galleguillos Madrid, F.M.; León, Y. Synthesis, crystal structures, Hirshfeld surface analysis, computational investigations, thermal properties, and electrochemical analysis of two new Cu(II) and Co(II) coordination polymers with the ligand 5-methyl-1-(pyridine-4-yl-methyl)-1H-1,2,3-triazole-4-carboxylate. Int. J. Mol. Sci. 2025, 26, 1671. [Google Scholar] [CrossRef] [Scilit]
  21. Bui, H.T.; Shrestha, N.K.; Khadtare, S.; Bathula, C.D.; Giebeler, L.; Noh, Y.; Han, S. Anodically Grown Binder-Free Nickel Hexacyanoferrate Film: Toward Efficient Water Reduction and Hexacyanoferrate Film Based Full Device for Overall Water Splitting. ACS Appl. Mater. Interfaces 2017, 9, 18015–18021. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  22. Kumar, Y.A.; Sana, S.S.; Ramachandran, T.; Assiri, M.A.; Rao, S.S.; Kim, S.C. From lab to field: Prussian blue frameworks as sustainable cathode materials. Dalton Trans. 2024, 53, 10770–10804. [Google Scholar] [CrossRef] [Scilit]
  23. Xi, Y.; Xiang, Y.; Zhang, C.; Bao, T.; Zou, Y.; Zou, J.; Wei, G.; Wang, L.; Xu, H.; Yu, C.; et al. Perfect Is Perfect: Nickel Prussian Blue Analogue as A High-Efficiency Electrocatalyst for Hydrogen Peroxide Production. Angew. Chem. Int. Ed. 2025, 64, e202413866. [Google Scholar] [CrossRef] [Scilit]
  24. Wang, Y.; Ding, H.; Deng, C.; Huang, T.; Xiao, J.; Xie, M.; Zhang, L.; Xu, G. Research Progress and Prospects of Prussian Blue Analogs and Their Derivatives in Small-Molecule Oxidative Coupled Hydrogen Production. Chem. Asian J. 2025, 20, e00665. [Google Scholar] [CrossRef] [Scilit]
  25. Liu, X.; Chi, J.; Mao, H.; Wang, L. Principles of Designing Electrocatalyst to Boost Reactivity for Seawater Splitting. Adv. Energy Mater. 2023, 13, 2301438. [Google Scholar] [CrossRef] [Scilit]
  26. Jiménez-Arévalo, V.M.; Martin, P.; Sepúlveda, M.F.; Azocar, M.I.; Zhou, X.; Madrid, F.M.G.; Ramirez, C.; Zagal, J.H.; Páez, M. Recent advances in high-entropy alloys for electrocatalysis: From rational design to functional performance. Mater. Des. 2025, 258, 114633. [Google Scholar] [CrossRef] [Scilit]
  27. Pashakolaie, V.G.; Gudlaugsson, B.; Ahmed, T.G. From Investment to Impact: Exploring Socio-Economic Prospect of Hydrogen Investment in Tees Valley, UK. Socio-Econ. Plan. Sci. 2026, 103, 102371. [Google Scholar] [CrossRef] [Scilit]
  28. Kyrimis, S.; Dimas, P.; Stamopoulos, D.; Tsakanikas, A. Socio-Economic Impact Assessment of Hydrogen Injection in the Natural Gas Network †. Energies 2025, 18, 725. [Google Scholar] [CrossRef] [Scilit]
  29. PAmoatey, P.; Al-Nadabi, A.; Chen, M.; Nikoo, M.R.; Al-Maktoumi, A.; Al-Hinai, A.; Izady, A. Transition to sustainable hydrogen energy in Oman: Implication for future socio-economic transformation and environmental well-being. Environ. Dev. Sustain. 2024, 27, 15213–15234. [Google Scholar] [CrossRef] [Scilit]
  30. Delgado, G.E.; Cisterna, J.; Llanos, J.; Pulido, R.; Naveas, N.; Narea, P.; Amo-Ochoa, P.; Zamora, F.; León, Y.; Brito, I. Structure–Property Relationships in Zwitterionic Pyridinium–Triazole Ligands: Insights from Crystal Engineering and Hirshfeld Surface Analysis. Int. J. Mol. Sci. 2025, 26, 5123. [Google Scholar] [CrossRef] [Scilit]
  31. Kjeldgaard, S.; Wagemaker, M.; Iversen, B.B.; Bentien, A. Sample dependent performance of aqueous copper hexacyanoferrate/zinc batteries. Mater. Adv. 2021, 2, 2036–2044. [Google Scholar] [CrossRef] [Scilit]
  32. Zampardi, G.; Warnecke, M.; Tribbia, M.; Glenneberg, J.; Santos, C.; La Mantia, F. Effect of the reactants concentration on the synthesis and cycle life of copper hexacyanoferrate for aqueous Zn-ion batteries. Electrochem. Commun. 2021, 126, 107030. [Google Scholar] [CrossRef] [Scilit]
  33. Palacios-Corella, M.; Echevarria, I.; Santos, C.S.; Schuhmann, W.; Ventosa, E.; Ibáñez, M. Prussian Blue Analogues as Anode Materials for Battery Applications: Complexities and Horizons. Chem. Mater. 2025, 37, 4203–4226. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  34. Polo-Cuadrado, E.; Osorio, E.; Acosta-Quiroga, K.; Camargo-Ayala, P.A.; Brito, I.; Rodriguez, J.; Alderete, J.B.; Forero-Doria, O.; Blanco-Acuña, E.F.; Gutiérrez, M. Nonlinear optical and spectroscopic properties, thermal analysis, and hemolytic capacity evaluation of quinoline-1,3-benzodioxole chalcone†. RSC Adv. 2024, 14, 10199–10208. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  35. Cabeza, L.F.; Martínez, F.R.; Borri, E.; Ushak, S.; Prieto, C. Thermal Energy Storage Using Phase Change Materials in High-Temperature Industrial Applications: Multi-Criteria Selection of the Adequate Material. Materials 2024, 17, 1878. [Google Scholar] [CrossRef] [Scilit]
  36. Yao, H.; Ni, L.; Wu, P.; Jiang, J.; Ni, Y.; Yao, X. Thermal hazard and pyrolysis mechanism of tetrazolo[1,5-a]pyridine by TG, DSC, ARC, TG-MS and DFT methods. J. Anal. Appl. Pyrolysis 2021, 159, 105299. [Google Scholar] [CrossRef] [Scilit]
  37. Concina, I. An Old Material for a New World: Prussian Blue and Its Analogues as Catalysts for Modern Needs. Inorganics 2024, 12, 124. [Google Scholar] [CrossRef] [Scilit]
  38. Wessells, C.D.; Huggins, R.A.; Cui, Y. Copper hexacyanoferrate battery electrodes with long cycle life and high power. Nat. Commun. 2011, 2, 550. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  39. Madrid, F.M.G.; Soliz, A.; Cáceres, L.; Salazar-Avalos, S.; Guzmán, D.; Gálvez, E. Corrosion of Reinforced A630-420H Steel in Direct Contact with NaCl Solution. Materials 2023, 16, 6017. [Google Scholar] [CrossRef] [Scilit]
  40. Madrid, F.M.G.; Arancibia-Bravo, M.; Cisterna, J.; Soliz, Á.; Salazar-Avalos, S.; Guevara, B.; Sepúlveda, F.; Cáceres, L. Corrosion of Titanium Electrode Used for Solar Saline Electroflotation. Materials 2023, 16, 3514. [Google Scholar] [CrossRef] [Scilit]
  41. Soliz, A.; Guzmán, D.; Cáceres, L.; Madrid, F.M.G. Electrochemical Kinetic Analysis of Carbon Steel Powders Produced by High-Energy Ball Milling. Metals 2022, 12, 665. [Google Scholar] [CrossRef] [Scilit]
  42. Wagner, C. On the interpretation of corrosion processes through the superposition of electrochemical sub-processes and on the potential formation at mixed electrodes. Z. Elektrochem. Angew. Phys. Chem. 1938, 44, 391–402. [Google Scholar] [CrossRef] [Scilit]
  43. Lukács, Z.; Kristóf, T. Linear transformations of the Butler–Volmer equation. Electrochem. Commun. 2023, 154, 107556. [Google Scholar] [CrossRef] [Scilit]
  44. Soliz, A.; Cáceres, L. Corrosion Behavior of Carbon Steel in LiBr in Comparison to NaCl Solutions under Controlled Hydrodynamic Conditions. Int. J. Electrochem. Sci. 2015, 10, 5673–5693. [Google Scholar] [CrossRef] [Scilit]
  45. Cisterna, J.; Araneda, C.; Narea, P.; Cárdenas, A.; Llanos, J.; Brito, I. The positional isomeric effect on the structural diversity of Cd(II) coordination polymers, using flexible positional isomeric ligands containing pyridyl, triazole, and carboxylate fragments. Molecules 2018, 23, 2634. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  46. Al-Matarneh, M.C.; Nicolescu, A.; Dascalu, I.A.; Shova, S.; Varganici, C.D.; Fifere, A.; Danac, R.; Marinas, I.C. Synthesis of New Zinc and Copper Coordination Polymers Derived from Bis (Triazole) Ligands. Crystals 2024, 14, 144. [Google Scholar] [CrossRef] [Scilit]
  47. Yang, F.; Xu, Y.; Wang, P.; Lin, Q.; Lu, M. Oxygen-Enriched Metal-Organic Frameworks Based on 1-(Trinitromethyl)-1 H-1,2,4-Triazole-3-Carboxylic Acid and Their Thermal Decomposition and Effects on the Decomposition of Ammonium Perchlorate. ACS Appl. Mater. Interfaces 2021, 13, 21516–21526. [Google Scholar] [CrossRef] [Scilit]
  48. Baghodrat, M.; Zampardi, G.; Glenneberg, J.; La Mantia, F. Influence of the Thermal Treatment on the Structure and Cycle Life of Copper Hexacyanoferrate for Aqueous Zinc-Ion Batteries. Batteries 2023, 9, 170. [Google Scholar] [CrossRef] [Scilit]
  49. Parajuli, D.; Tanaka, H.; Sakurai, K.; Hakuta, Y.; Kawamoto, T. Thermal decomposition behavior of prussian blue in various conditions. Materials 2021, 14, 1151. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  50. Moloney, M.P.; Massoni, N.; Grandjean, A. Tuning the thermal stability of copper(II) hexacyanoferrate(II) nanoparticles. J. Therm. Anal. Calorim. 2021, 145, 2353–2362. [Google Scholar] [CrossRef] [Scilit]
  51. Guadagnini, L.; Tonelli, D.; Giorgetti, M. Improved performances of electrodes based on Cu2+-loaded copper hexacyanoferrate for hydrogen peroxide detection. Electrochim. Acta 2010, 55, 5036–5039. [Google Scholar] [CrossRef] [Scilit]
  52. Wang, P.; Wang, P.; Wu, T.; Zhao, C.; Pu, Z.; Zhang, Y. Interface Engineering of NiMoSx@NiMnFe Prussian Blue Analogue Nanowires to Efficiently Boost Overall Seawater Splitting at High Current Densities. Adv. Funct. Mater. 2025, 35, 2417924. [Google Scholar] [CrossRef] [Scilit]
  53. Xiao, Z.; Zhu, X.; Bai, L.; Liu, Z. Porous Prussian Blue Analogs Decorated Electrospun Carbon Nanofibers as Efficient Electrocatalyst for Overall Water Splitting. Energies 2024, 17, 1154. [Google Scholar] [CrossRef] [Scilit]
  54. Madrid, F.M.G.; Arancibia-Bravo, M.P.; Sepúlveda, F.D.; Lucay, F.A.; Soliz, A.; Cáceres, L. Ultrafine Kaolinite Removal in Recycled Water from the Overflow of Thickener Using Electroflotation: A Novel Application of Saline Water Splitting in Mineral Processing. Molecules 2023, 28, 3954. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  55. Soliz, A.; Madrid, F.M.G.; Murcia, J.A.C.; Angulo, S.; Avalos, S.S.; Fariñas, B.A.; Guzmán, A. Electrochemical Performance of Ti Gr. 2 as Electrodes in Contact with Saline Suspension of Clays during the Electroflotation Process. Appl. Sci. 2024, 14, 8825. [Google Scholar] [CrossRef] [Scilit]
  56. Abhilash, S.; Sumi, V.S.; Sarika, S.; Deepa, J.P.; Sreekala, C.O.; Rijith, S. Development and characterization of NiMoP alloy coating for electrocatalytic hydrogen evolution reaction in alkaline media. Fuel 2022, 318, 123598. [Google Scholar] [CrossRef] [Scilit]
  57. Van Ede, M.C.; Angst, U. Tafel slopes and exchange current densities of oxygen reduction and hydrogen evolution on steel: Influence of the electrochemical measurement procedure. Corros. Eng. Sci. Technol. 2023, 59, 39–55. [Google Scholar] [CrossRef] [Scilit]
  58. Bi, S.; Geng, Z.; Zhao, L.; Gao, Z.; Qu, C.; Li, W.; Jin, L.; Xue, M.; Zhang, C. Porous Nickel-Molybdenum Phosphide Alloy Electrodes for Alkaline Hydrogen Evolution Reaction at the High-Current Density. Renewables 2024, 2, 150–161. [Google Scholar] [CrossRef] [Scilit]
  59. Wang, X.; Xiao, X.; Chen, C.; Hu, J.; Zhang, L.; Sun, D.; Xu, X. The interfacial engineering of seaurchin-like CoP/Ni2P heterostructure for highly efficient alkaline hydrogen evolution. Chem. Eng. Sci. 2025, 301, 120740. [Google Scholar] [CrossRef] [Scilit]
  60. Zhang, H.; Li, Z.; Qiao, L.; Chi, K.; Guo, X.; Cao, D.; Cheng, D. Constructing CoP/Ni2P heterostructure confined Ru sub-nanoclusters for enhanced water splitting in wide pH conditions. Adv. Sci. 2024, 11, 2401398. [Google Scholar] [CrossRef] [Scilit]
  61. El Hanafi, M.; BaQais, A.A.; Saadi, M.; Ahsaine, H.A. Advances and Outlook of Metal–Organic Frameworks as High-Performance Electrocatalysts for Hydrogen Evolution Reaction: A Minireview. Energy Fuels 2023, 37, 10869–10885. [Google Scholar] [CrossRef] [Scilit]
  62. Nemiwal, M.; Gosu, V.; Zhang, T.C.; Kumar, D. Metal organic frameworks as electrocatalysts: Hydrogen evolution reactions and overall water splitting. Int. J. Hydrogen Energy 2021, 46, 10216–10238. [Google Scholar] [CrossRef] [Scilit]
  63. Yang, L.J.; Guan, H.Y.; Yuan, S.; Sun, T.; Jiang, A.N.; Feng, J.J. Research progress of chlorine corrosion resistance in seawater electrolysis: Materials and technologies. Chem. Eng. J. 2024, 503, 158458. [Google Scholar] [CrossRef] [Scilit]
  64. Mohammadnezhad, G.; Amirian, A.M.; Gorls, H.; Plass, W.; Sandleben, A.; Schafer, S.; Klein, A. Redox Instability of Copper(II) Complexes of a Triazine-Based PNP Pincer. Eur. J. Inorg. Chem. 2021, 2021, 1140–1151. [Google Scholar] [CrossRef] [Scilit]
  65. Le Fur, M.; Beyler, M.; Le Poul, N.; Lima, L.M.P.; Le Mest, Y.; Delgado, R.; Iglesias, C.P.; Patinec, V.; Tripier, R. Improving the stability and inertness of Cu(ii) and Cu(i) complexes with methylthiazolyl ligands by tuning the macrocyclic structure. Dalton Trans. 2016, 45, 7406–7420. [Google Scholar] [CrossRef] [Scilit]
  66. Foppa, L.; Ruther, F.; Geske, M.; Koch, G.; Girgsdies, F.; Kube, P.; Carey, S.J.; Havecker, M.; Timpe, O.; Tarasov, A.V.; et al. Data-Centric Heterogeneous Catalysis: Identifying Rules and Materials Genes of Alkane Selective Oxidation. J. Am. Chem. Soc. 2023, 145, 3427–3442. [Google Scholar] [CrossRef] [Scilit]
  67. Loboda, O.; Strizhak, P.; Góra, R.W.; Cervinka, C. Editorial: Fragment-based electronic structure methods for solids. Front. Chem. 2023, 11, 1165201. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  68. Xu, J.; Lund, C.; Patel, P.; Kim, Y.L.; Liu, C. Recent Advances on Computational Modeling of Supported Single-Atom and Cluster Catalysts: Characterization, Catalyst–Support Interaction, and Active Site Heterogeneity. Catalysts 2024, 14, 224. [Google Scholar] [CrossRef] [Scilit]
  69. Souza, R.A.C.; Costa, W.R.P.; Faria, E.F.; Bessa, M.A.S.; Menezes, R.P.; Martins, C.H.G.; Maia, P.I.S.; Deflon, V.M.; Oliveira, C.G. Copper(II) complexes based on thiosemicarbazone ligand: Preparation, crystal structure, Hirshfeld surface, energy framework, antiMycobacterium activity, in silico and molecular docking studies. J. Inorg. Biochem. 2021, 223, 111543. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  70. MacK, K.; Leupoldt, A.W.V.; Forster, C.; Ezhevskaya, M.; Hinderberger, D.; Klinkhammer, K.W.; Heinze, K. Effect of chelate ring expansion on Jahn-Teller distortion and Jahn-Teller dynamics in copper(II) complexes. Inorg. Chem. 2012, 51, 7851–7858. [Google Scholar] [CrossRef] [Scilit]
  71. Deeth, R.J.; Hearnshaw, L.J.A. Molecular modelling of Jahn–Teller distortions in Cu(ii)N6complexes: Elongations, compressions and the pathways in between. J. Chem. Soc. Dalton Trans. 2005, 6, 1092–1100. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  72. Anantharaj, S.; Noda, S.; Jothi, V.R.; Yi, S.C.; Driess, M.; Menezes, P.W. Strategies and Perspectives to Catch the Missing Pieces in Energy-Efficient Hydrogen Evolution Reaction in Alkaline Media. Angew. Chem. Int. Ed. Engl. 2021, 60, 18981–19006. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  73. Velez, J.H.; Aguirre, M.J.; Zamora, P.P.; Cattin, M.B.L.; Makha, M.; Bernede, J.C. Fabrication Parameter Optimization for a Multilayer Photovoltaic Cell Based on the Heterojunction: Zinc(II)-Meso-Tetrakis(4-Bromophenyl) Porphyrins/Fullerenes. Open J. Appl. Sci. 2013, 3, 136–144. [Google Scholar] [CrossRef]
  74. Cárdenas, C.; Rabi, N.; Ayers, P.W.; Morell, C.; Jaramillo, P.; Fuentealba, P. Chemical reactivity descriptors for ambiphilic reagents: Dual descriptor, local hypersoftness, and electrostatic potential. J. Phys. Chem. A 2009, 113, 8660–8667. [Google Scholar] [CrossRef] [Scilit]
  75. Del Valle, M.A.; Gacitúa, M.A.; Borrego, E.D.; Zamora, P.P.; Díaz, F.R.; Camarada, M.B.; Antilén, M.P.; Soto, J.P. Electro-Synthesis and Characterization of Aniline and O-Anisidine Oligomers. Int. J. Electrochem. Sci. 2012, 7, 2552–2565. [Google Scholar] [CrossRef] [Scilit]
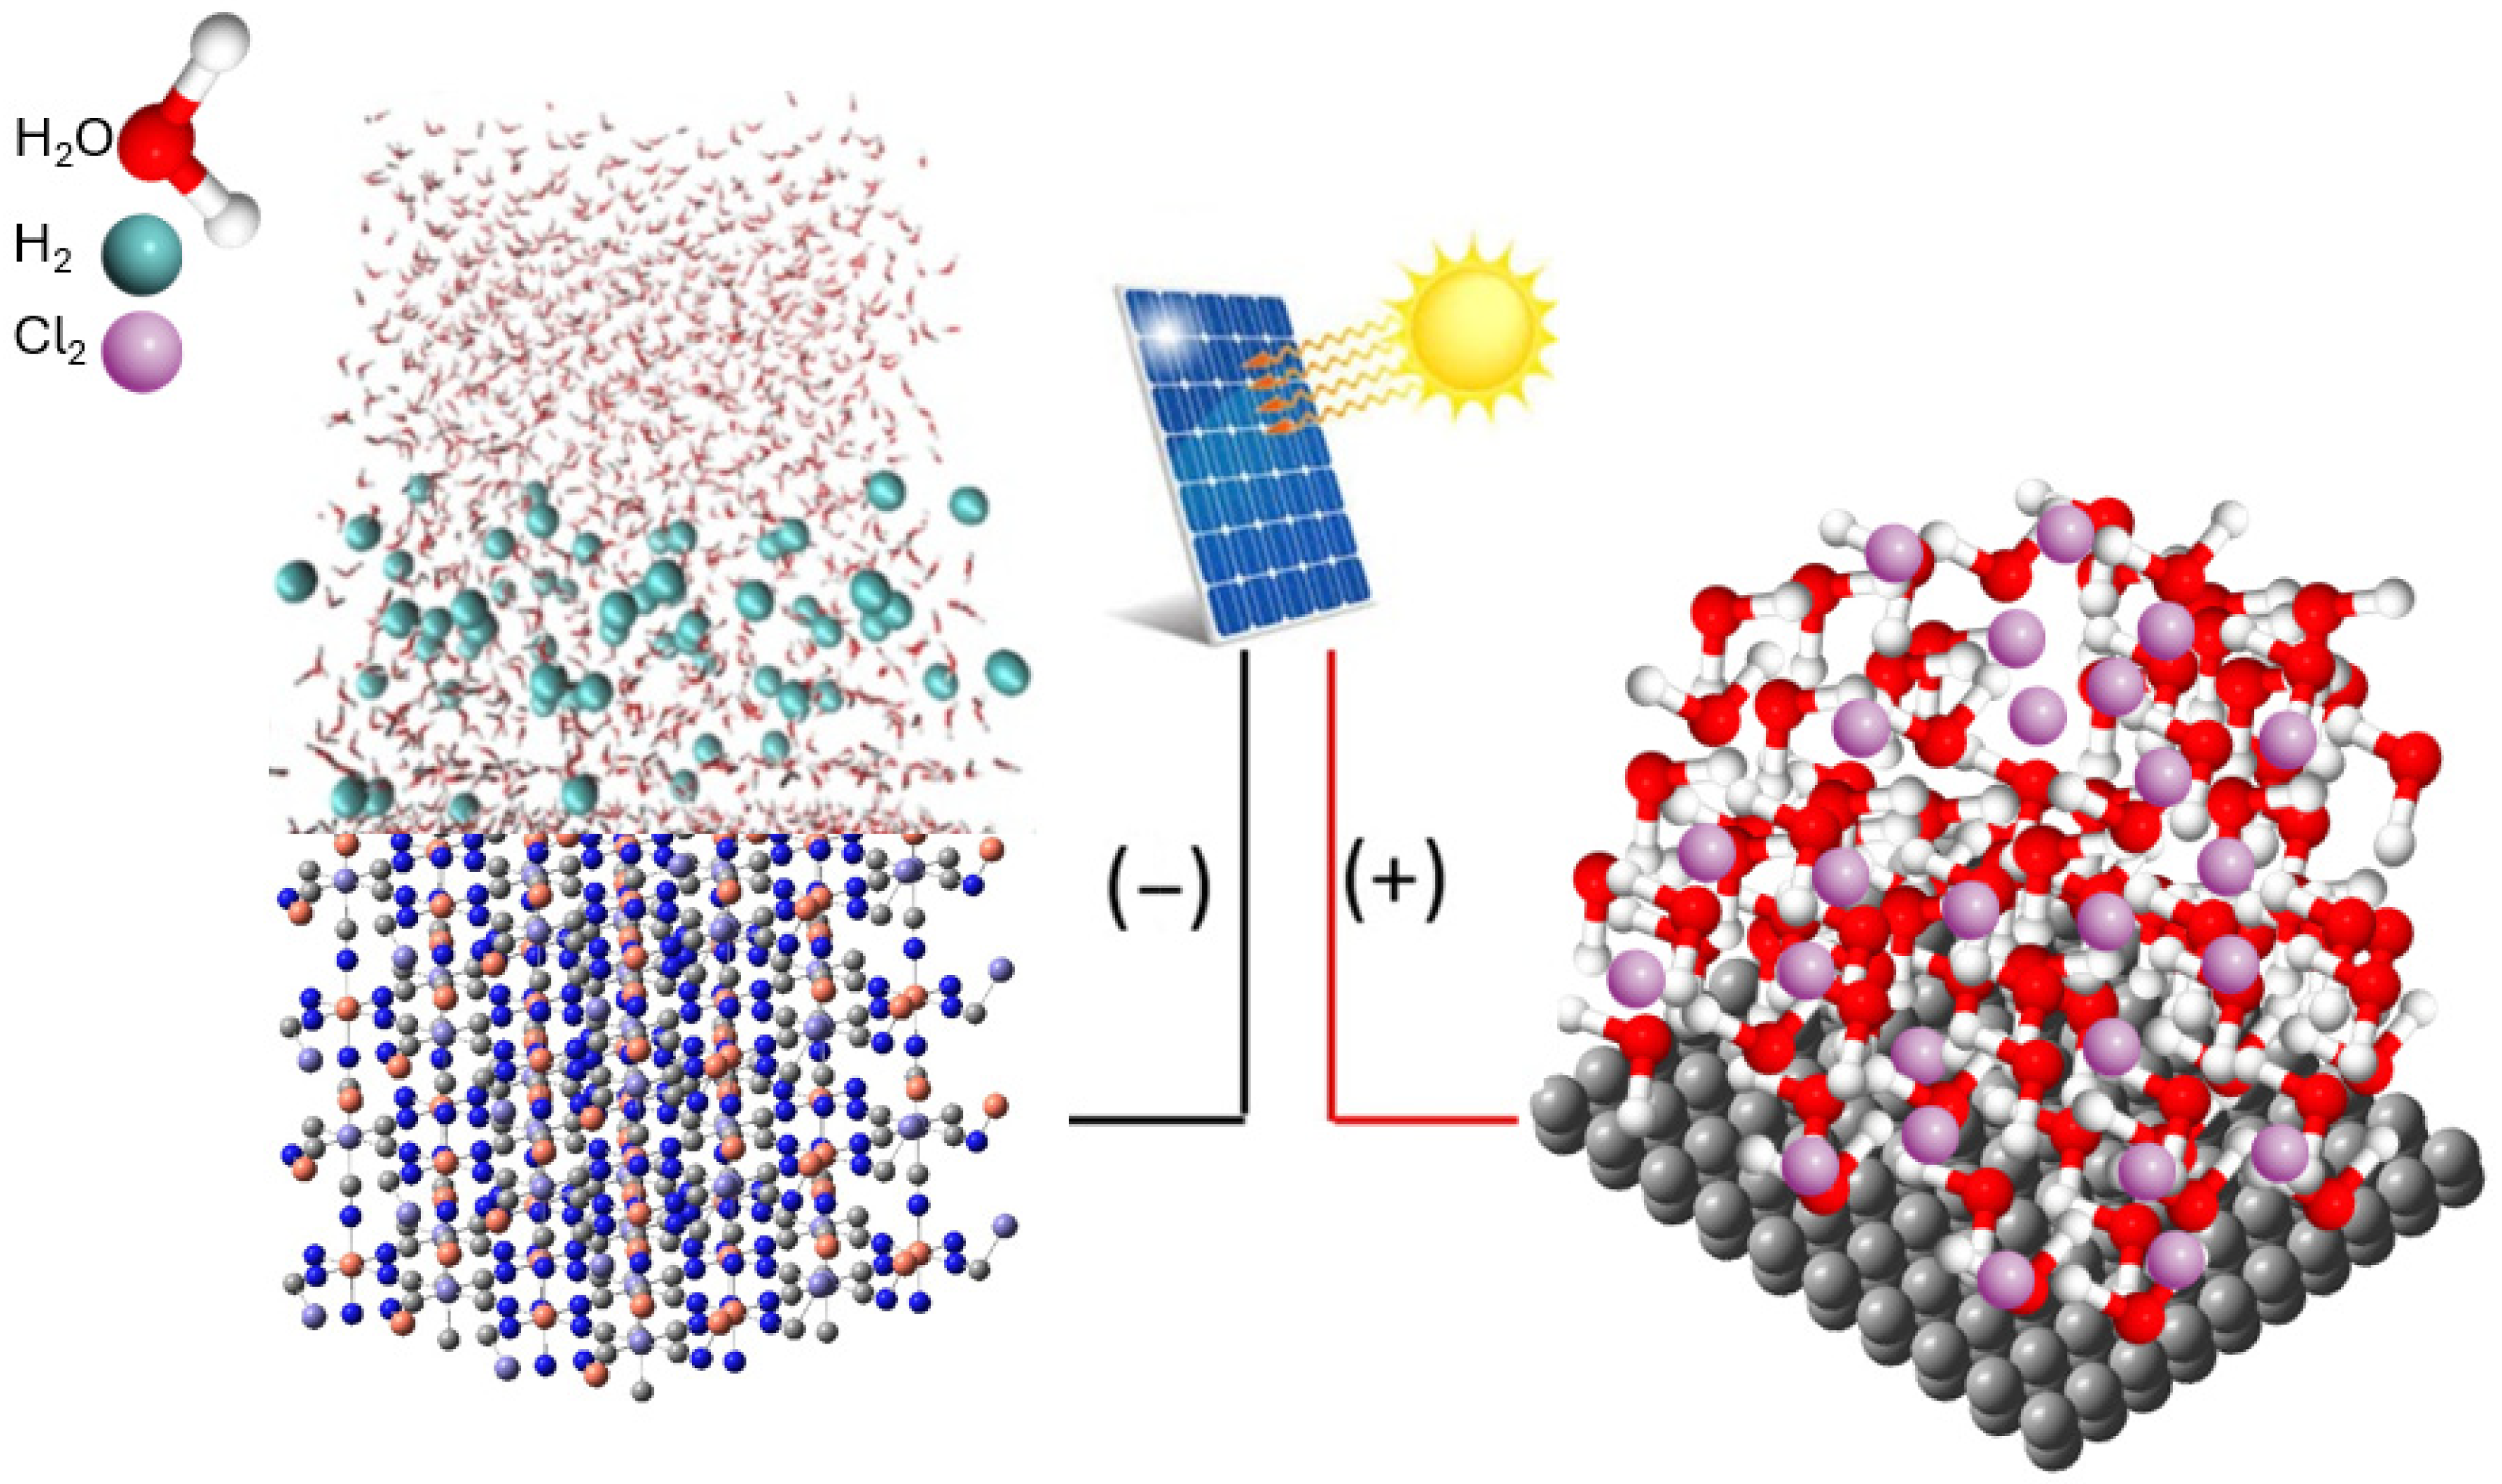
Figure 1. Schematic illustration of CuHCF as a cathode electrode for hydrogen and other metals for chlorine generation. Orange Ball = Copper, Blue Ball = Nitrogen, and Gray Ball = Carbon.
Processes 14 00423 g001
Figure 2. Synthesis of a triazole–carboxylate ligand (L1) derived from 3-azidopyridine, proceeding first through an ester intermediate and finally transforming it into its sodium carboxylate salt.
Processes 14 00423 g002
Figure 3. (a) FT-IR spectra of the organic ligand precursor (L1) and its corresponding coordination polymer (CCCu); and (b) FT-IR spectra of the ligand [K3(Fe(CN)6)] (L2) and its corresponding coordination polymer of CuHCF, and (c) Table summary of characteristic FT-IR vibrational bands and assignments (L1, CCCu, L2, CuHCF).
Processes 14 00423 g003
Figure 4. Two-dimensional chemical structures of (a) CCCu (copper coordination compound) and (b) CuHCF (copper hexacyanoferrate), respectively.
Processes 14 00423 g004
Figure 5. (a) Thermogravimetric (TG) and differential scanning calorimetry (DSC) curves of the coordination polymer CCCu. (b) Thermogravimetric (TG) and differential scanning calorimetry (DSC) curves of the CuHCF compound.
Processes 14 00423 g005
Figure 6. Three-dimensional model of the molecular structure of CCCu. Image obtained from our previous work (adapted from [20]). Red Ball = Oxygen, Orange Ball = Copper, Blue Ball = Nitrogen, Gray Ball = Carbon.
Processes 14 00423 g006
Figure 7. (a) DRX from CCCu [20], and (b) the CCCu molecule. Red Ball = Oxygen, Orange Ball = Copper, blue Ball = Nitrogen, Gray Ball = Carbon.
Processes 14 00423 g007
Figure 8. (a) X-ray diffraction pattern of Cu1.333[Fe(CN)6]0.667·H2O (CuHCF), showing the peaks corresponding to the (200), (220), (400), (420), and (422) planes of the Fm3-m space group [4], and (b) schematic representation of the crystal structure of CuHCF, showing the A sites, Cu2+ ions, and Fe3+ ions. Orange Ball = Copper, blue Ball = Nitrogen, Gray Ball = Carbon.
Processes 14 00423 g008
Figure 9. SEM and EDS characterization of CCCu: (a) overall laminar morphology (20 µm); (b) mid-magnification detail (2 µm); (c) high-magnification view (500 nm); (d) global EDS spectrum; and (e) elemental maps (100 µm).
Processes 14 00423 g009
Figure 10. (a) general morphology showing aggregated spherical particles (b) morphological detail at intermediate magnification, highlighting surface texture and interparticle porosity (c) high-resolution image revealing nanostructured features on particle surfaces (d) global energy-dispersive X-ray (EDS) spectrum indicating elemental composition; (e) elemental distribution maps for C, N, Cu, Fe, O, and K.
Processes 14 00423 g010
Figure 11. (a,b) Potentiodynamic polarization curve and superposition model output curves for materials for CCCu and CuHCF, (c,d) Tafel diagram for CCCu and CuHCF in aerated 0.5 M NaCl and 2 mV/s.
Processes 14 00423 g011
Figure 12. Hydrogen production mechanisms in CuHFC steps 1 and 2 are non-catalytic.
Processes 14 00423 g012
Figure 13. Selected section for mechanistic and stability study: (a) front view of the crystal; (b) side view of the crystal. Orange Ball = Copper, blue Ball = Nitrogen, Gray Ball = Carbon and Purple Ball: Sodium.
Processes 14 00423 g013
Figure 14. Gibbs free energy IRC for hydrogen production on CuHCF.
Processes 14 00423 g014
Figure 15. OPDOS analysis of the CuHCF-N complex.
Processes 14 00423 g015
Figure 16. RDG analysis of the CuHCF complex in saline medium: (a) pre-reaction state (step 1); (b) intermediate after water incorporation (step 3); (c) Cu+ state prior to hydrogen evolution (step 4), (d) RDG TS1 (e) TS2 respectively. Negative frequencies are found in Table S1 (Supplementary Materials).
Processes 14 00423 g016
Figure 17. Hydrogen production mechanism in CCCu; steps 1 and 2 are non-catalytic.
Processes 14 00423 g017
Figure 18. Fragment of the CCCu complex chosen for mechanistic and stability analysis. Red Ball = Oxygen, Orange Ball = Copper, Blue Ball = Nitrogen, and Gray Ball = Carbon.
Processes 14 00423 g018
Figure 19. Gibbs free energy IRC for hydrogen production on CCCu.
Processes 14 00423 g019
Figure 20. OPDOS of the CCCu complex: (a) Cu–N interactions; (b) Cu–O interactions.
Processes 14 00423 g020
Figure 21. RDG iso-surface of the CCCu complex in the Cu2+ intermediate state with coordinated OH−. (a) CCCu complex before the reaction, (b) stabilization of the intermediate via hydrogen-bond bridging, (c) stabilization of the intermediate by carboxyl groups and sodium atoms, (d) TS1 CCCu, and (e) TS2 CCCu. Negative frequencies are found in Table S1 (Supplementary Materials).
Processes 14 00423 g021
Table 1. Data obtained from the superposition model based on mixed potential theory.
ParametersCCCuCuHCF
i0,H2, A/m2−0.062−0.019
tH2, mV/dec−301−222
i0,O2, A/m2−2.251·10−6−7.681·10−7
iL,O2, A/m2−1.279−1.362
tO2, mV/dec−157−238
i0,M, A/m20.3950.74
tM, mV/dec176353
Ecorr, mV/SHE−23−40
icorr, A/m20.3790.259
Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content.