Skip to Content
  • 2.4
    Impact Factor
  • 41 days
    Submission to First Decision
  • 9 days
    Acceptance to Publication

Thalassemia Reports

Thalassemia Reports is an international, peer-reviewed, open access journal on the study, diagnosis, and treatment of thalassemia, published quarterly online by MDPI (from Volume 12, Issue 1 - 2022).

Get Alerted

Add your email address to receive forthcoming issues of this journal.

All Articles (299)

  • Article
  • Open Access

Background/Objectives: Beta-thalassemia and sickle cell disorders are among the most common inherited hemoglobin disorders affecting tribal populations in India, with marked ethnic and geographic heterogeneity. Although exploratory multivariate approaches have been used, only limited evidence is available regarding the distribution and clustering patterns of these disorders among Scheduled Tribe populations in Tamil Nadu. This study aimed to examine the distribution and population-specific clustering of beta-thalassemia- and sickle cell-related disorders among hematologically confirmed cases of the scheduled tribes from the selected districts. Methods: A cross-sectional analytical study was conducted among the Scheduled Tribe populations in the selected districts of Tamil Nadu. Confirmed cases of hemoglobin disorders were identified through complete blood count, peripheral smear examination, reticulocyte count, and high-performance liquid chromatography (HPLC). A total of 89 confirmed cases were included in the exploratory analysis. Disease forms were categorised as beta (β)-thalassemia related disorders and sickle cell- related disorders, including sickle cell β-thalassemia. Chi-square tests were performed to assess associations with socio-demographic and environmental variables. Correspondence analysis and multidimensional scaling were applied to identify clustering patterns and structural relationships. Results: Of the 89 confirmed cases, 48 (53.9%) had β-thalassemia and related disorders (predominantly beta-thalassemia minor), while 41 (46.1%) had sickle cell-related and combined forms (predominantly sickle cell trait). There were significant associations between disease forms and tribal affiliation (p < 0.001), drinking water source (p < 0.001), and altitude of residence (p = 0.003). Correspondence analysis demonstrated distinct clustering of β-thalassemia- and sickle cell-related disorders across tribes and districts. Multidimensional scaling further indicated that population structure and geographic context were more strongly associated with disease distribution than individual demographic variables. Conclusions: Hemoglobinopathy disease forms among Scheduled Tribes in the selected districts of Tamil Nadu showed clear eco-geographical and population-specific clustering patterns. The combination of correspondence analysis and multidimensional scaling revealed latent clustering patterns beyond those identified using traditional statistical analyses, emphasizing the intricate relationship between mutation distribution, environment, and population structure. These results highlight the significance of tribe-specific, equity-focused screening and genetic counselling programmes. To close diagnostic gaps and improve precision-based approaches in underserved populations, public health interventions must be tailored to intra-tribal heterogeneity. These findings may help inform future studies on targeted, population-specific screening and decentralised diagnostic strategies to reduce health inequities in the selected Scheduled Tribe populations.

Thalass. Rep.

8 October 2026
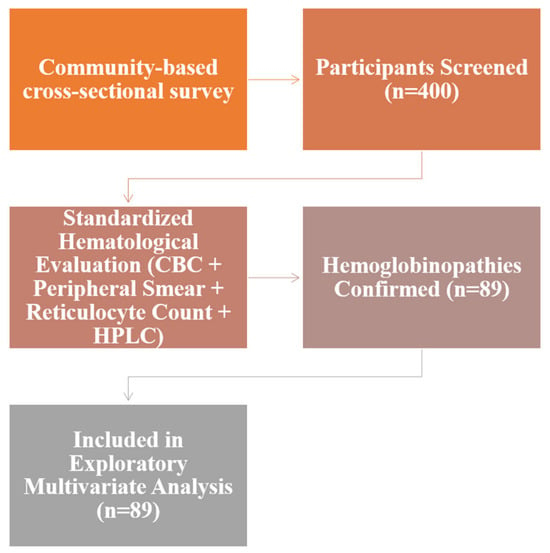
Flow diagram of participant selection for the exploratory multivariate analysis.
  • Article
  • Open Access

Background/Objectives: Severe unstable hemoglobinopathies may cause transfusion dependence from early childhood, and allogeneic hematopoietic cell transplantation (allo-HCT) is the only established curative option. Pediatric experience with allo-HCT for unstable hemoglobinopathies is limited to isolated reports, and long-term outcomes remain poorly characterized. The objective of this study was to characterize the clinical and molecular features, transplant approaches, and long-term outcomes of two Brazilian patients with unstable hemoglobinopathies, in the context of the published pediatric literature. Methods: We performed a retrospective analysis of two children with unstable hemoglobinopathies who underwent allo-HCT at a Brazilian center. PubMed/MEDLINE and Scopus were searched for pediatric cases published between 2002 and 2025 using terms related to unstable hemoglobin variants and hematopoietic transplantation. Published cases were combined with the Brazilian patients to form an aggregated cohort. Clinical, molecular, transplant, and follow-up data were analyzed descriptively. Results: The aggregated cohort included 13 patients and 15 allo-HCT procedures, using matched sibling, haploidentical, and other alternative donors. All patients were transfusion-dependent before transplantation. Two patients required retransplantation after graft failure. At last follow-up, the nine published patients with available outcome data and both Brazilian patients were alive and transfusion independent. The Brazilian patients maintained complete donor chimerism and transfusion independence for 8 and 9 years, the longest follow-up reported in this population. One remained transfusion independent after retransplantation for secondary graft failure. Conclusions: Allo-HCT may achieve durable transfusion independence in children with severe unstable hemoglobinopathies, but the small number of reported cases limits the interpretation of these findings. Our series adds long-term follow-up data supporting this approach and highlights the need for larger, prospective studies to guide donor selection and conditioning strategies in this rare population.

Thalass. Rep.

8 October 2026

  • Review
  • Open Access

Background: Sickle cell disease (SCD) and β-thalassemia are chronic thromboinflammatory disorders associated with venous, arterial, and microvascular thrombotic complications. However, evidence regarding disease-specific risk stratification and thrombosis prevention remains fragmented. This review summarizes current evidence on the mechanisms, risk identification, and prevention of thrombosis in these disorders. Methods: A narrative review of the literature was conducted using PubMed and Scopus to identify studies evaluating the thrombotic mechanisms, risk factors, biomarkers, and preventive strategies in SCD and β-thalassemia. Results: Thrombotic complications arise through interconnected mechanisms involving chronic hemolysis, nitric oxide depletion, endothelial dysfunction, phosphatidylserine exposure, microparticle generation, platelet activation, tissue factor expression, and enhanced thrombin generation. ADAMTS13–von Willebrand factor dysregulation may contribute to platelet adhesion and microvascular thrombosis, whereas NETosis and complement activation remain emerging mechanisms requiring further clinical validation. In SCD, thrombotic risk is associated with factors including pregnancy, recurrent hospitalization, central venous access devices, prior thrombosis, and pulmonary hypertension. In β-thalassemia, higher thrombotic event rates have been reported in non-transfusion-dependent disease, particularly among patients who have undergone splenectomy or have thrombocytosis, severe anemia, nucleated erythrocytosis, or pulmonary hypertension. Although several biomarkers reflect thromboinflammatory activity, none has been prospectively validated for routine risk prediction. Thromboprophylaxis is therefore generally limited to established high-risk clinical settings and is largely extrapolated from general venous thromboembolism guidelines. Observational data suggest that direct oral anticoagulants may be associated with similar rates of VTE recurrence and possibly lower rates of major bleeding than vitamin K antagonists in SCD; however, the certainty of evidence is very low. Conclusions: Current thrombosis management in SCD and β-thalassemia relies primarily on individualized risk assessment and extrapolation from general VTE recommendations. Prospective multicenter studies, randomized trials, and validated disease-specific risk prediction models are needed to optimize thrombosis prevention and anticoagulation strategies in these disorders.

Thalass. Rep.

1 October 2026

  • Review
  • Open Access

Background/Objectives: Endothelial dysfunction is increasingly recognized as a key contributor to vascular complications in β-thalassemia but remains insufficiently integrated into clinical assessment and therapeutic trial design. This review summarizes current evidence on the mechanisms, biomarkers, vascular imaging findings, and therapeutic perspectives of endothelial dysfunction across the clinical spectrum of β-thalassemia. Methods: A focused PubMed search was conducted using predefined terms related to endothelial dysfunction, nitric oxide, oxidative stress, adhesion molecules, extracellular vesicles, vascular imaging, and therapeutics in β-thalassemia. Original studies and systematic reviews evaluating endothelial dysfunction, vascular biomarkers, imaging, or cardiovascular outcomes in β-thalassemia were included. Results: Endothelial dysfunction arises from the interplay of hemolysis-driven nitric oxide depletion, free heme- and iron-mediated oxidative injury, chronic inflammation, extracellular vesicle-mediated vascular activation, and hypercoagulability. Meta-analyses demonstrate significant elevations in intercellular adhesion molecule-1, vascular cell adhesion molecule-1, E-selectin, P-selectin, and endothelin-1. Emerging biomarkers include endothelial activation and stress index, adipocytokines, and serum metabolomics. Vascular imaging demonstrates impaired flow-mediated dilation, increased carotid intima-media thickness, elevated pulse-wave velocity, and pulmonary hypertension, with distinct vascular phenotypes in transfusion-dependent thalassemia (TDT) and non-transfusion-dependent thalassemia (NTDT). Iron chelation, luspatercept, mitapivat, and curative gene-based therapies have strong biological rationale for improving endothelial function, although vascular endpoints have rarely been evaluated. Conclusions: Endothelial dysfunction is a multidimensional and potentially modifiable component of β-thalassemia vasculopathy. Future clinical trials should incorporate standardized endothelial biomarkers and vascular imaging as predefined endpoints and evaluate TDT and NTDT separately to advance mechanism-based vascular therapies.

Thalass. Rep.

7 September 2026

Featured Articles of Last Quarter

Highly Accessed Articles

News & Conferences

Latest Issues

Open for Submission

Journal Sections

XFacebookLinkedIn
Thalass. Rep. - ISSN 2039-4365