
Communication

Completion of the Genome Sequence of a Historic CDV Vaccine Strain, Rockborn: Evolutionary and Epidemiologic Implications

1
National Laboratory of Virology, Szentágothai Research Centre, University of Pécs, 7624 Pécs, Hungary
2
Faculty of Sciences, Institute of Biology, University of Pécs, 7624 Pécs, Hungary
3
Department of Medical Biology, Medical School, University of Pécs, 7624 Pécs, Hungary
4
Department of Pharmacology and Toxicology, University of Veterinary Medicine, 1078 Budapest, Hungary
5
National Laboratory for Infectious Animal Diseases, Antimicrobial Resistance, Veterinary Public Health and Food Chain Safety, HUN-REN Veterinary Medical Research Institute, Hungária krt. 21., 1077 Budapest, Hungary
6
Department of Veterinary Medicine, University of Bari Aldo Moro, 70010 Valenzano, Italy
*
Author to whom correspondence should be addressed.
These authors contributed equally to this work.
Vet. Sci. 2026, 13(1), 81; https://doi.org/10.3390/vetsci13010081
Submission received: 31 October 2025 / Revised: 8 January 2026 / Accepted: 11 January 2026 / Published: 14 January 2026
(This article belongs to the Section Veterinary Microbiology, Parasitology and Immunology)

Simple Summary

Canine distemper is a serious, contagious disease in dogs and other animals, and vaccines are crucial for its control. An old vaccine strain, called Rockborn, was widely used but later withdrawn from many markets due to safety concerns, although vaccines with Rockborn-like viruses are still on the market. We sequenced the entire genome of a Rockborn virus cell passage available in our laboratory to understand its evolution and improve future diagnostic tools. Our analyses revealed that viruses similar to the Rockborn strain are still circulating in wildlife and domestic dogs. More importantly, it was discovered that the Rockborn strain has been exchanging genetic material with other canine distemper viruses, creating new, recombinant viruses.

Abstract

The historic Rockborn strain of the canine distemper virus was widely used as a vaccine, but its use was discontinued due to safety concerns. Yet, Rockborn-like canine distemper virus strains are still used in some vaccine formulations. Genetic analysis of this strain was previously limited to the H gene, leaving its full evolutionary and pathogenic potential unclear. This study aimed to determine the complete genome sequence of the Rockborn strain to reconstruct its origin, understand its evolution, and provide a reference for improving diagnostics and future research on virulence markers. An amplicon-based sequencing protocol using MinION nanopore technology was employed to determine the complete genome of the Rockborn-46th laboratory strain. The genome was assembled, annotated, and analyzed in comparison with 223 genomes. The complete genome of the Rockborn strain was 15,690 nucleotides in length. Phylogenetic analysis revealed that Rockborn forms a unique lineage with field isolates from a masked civet in China and a dog in the United States. Crucially, a significant recombination event was identified, showing that the Rockborn strain acted as a parental strain, contributing its F and H genes to create mosaic viruses. The full-genome characterization of the Rockborn strain confirms that Rockborn-like viruses persist and actively contribute to the evolution of canine distemper virus through recombination. This finding highlights the inadequacy of single-gene analysis for diagnostics and surveillance, and underscores the necessity of whole-genome sequencing to accurately track the virus epidemiology and evolution.

1. Introduction

Canine distemper virus (CDV) is a paramyxovirus (Paramyxoviridae family, Morbillivirus genus) and possesses an enveloped virion with a minus-sense single-stranded RNA genome of ~15 kilobases in length. Among the six main primary gene products (N, P, M, F, H, and L) (Supplementary Table S1 [1,2,3,4,5,6,7,8,9,10,11,12]), the H gene is the most characterized genomic region. The H gene product acts as a cell attachment protein and mediates entry; moreover, antibodies against the H protein protect against infection and disease [13]. The exposure of the H protein to the immune system and the widespread use of CDV vaccines globally are thought to contribute to the genetic diversity among circulating strains. A genotyping classification scheme of CDV has been established based on the H gene [14,15,16,17].
The historic CDV strain, Rockborn, was isolated and attenuated by serial passage in the late 1950s [18]. Rockborn strain-based vaccines have been commercialized and distributed globally since the 1960s. This vaccine was shown to elicit a strong immune response and protect against disease caused by field CDV strains. However, safety concerns started to rise in the 1970s due to observed residual virulence. Although the etiological role was never fully proven, cases of vaccine-related encephalitis in the United States in the mid-1990s led to the definitive withdrawal of Rockborn-based CDV vaccines from the market [19,20]. Yet, several modern vaccines seemingly contain Rockborn-like viruses [21].
Overall, the high genetic identity between vaccine stock/laboratory strains and field isolates of vaccine-origin clinical cases was consistent with the hypothesis that the vaccine viruses retained residual virulence, likely in co-infection with other immunosuppressive pathogens. This was attributed to an intrinsic virulence of low-passage stocks established on primary dog kidney (DK) cells and grown for vaccine passage cultures on Madin Darby Canine Kidney (MDCK) cells. In most cases, increased virulence of the dog-kidney-cell-attenuated CDV was suspected to have been triggered by the contemporaneous presence in the vaccine formulations of live modified canine adenovirus type-1 [21]. Under experimental conditions, re-acquisition of virulence by the kidney-cell-attenuated Rockborn virus has been demonstrated in vivo after six sequential passages in dogs and in vitro after ten passages in primary dog lung macrophages, but the molecular bases of this phenotypic change were not investigated [22].
So far, genetic analysis of the Rockborn strain has been limited to the H gene. On sequence analysis of the full-length H gene of a cell passage (46th) of strain Rockborn, the virus and related strains were found to share 92–97% nucleotide (nt) and 91–96% amino acid (aa) sequence homology to other lineages of CDV. Additional partial H gene sequences were available for comparison, originating from vaccine-associated clinical cases and wild animals. Sequence divergence among strains varied slightly, with both nt and aa identities exceeding 99% [21]. With the determination of the full genome, we had the following objectives: (i) to better reconstruct the origin and evolution of the Rockborn strain and (ii) to extend the information with implications for diagnostic developments. Additionally, the reference genome sequence presented in this study will help identify virulence markers in clinically relevant Rockborn-associated CNS infections, if any.

2. Materials and Methods

2.1. Sequencing

The complete genome sequence of Rockborn-46th was determined using an amplicon-based sequencing protocol that combines amplification of adjacent CDV genomic fragments with MinION nanopore sequencing (Oxford Nanopore Technologies, Oxford, UK). The amplicon-based sequencing method for canine distemper virus, together with the primers employed, has been described previously [23], and the complete protocol is also available on the protocols.io page (https://www.protocols.io/view/universal-amplicon-based-sequencing-method-for-can-x54v9j6mpg3e/v1/metadata (accessed on 24 November 2021)). In brief, cDNA was prepared from extracted viral RNA with Superscript IV (Invitrogen, Carlsbad, CA, USA) using random hexamers. Overlapping PCR products were amplified from the cDNA with the Q5 Hot Start HF Polymerase (New England Biolabs, Ipswich, MA, USA) with multiple primer sets in parallel pools and then purified with AMPure XP beads (Beckman Coulter, Brea, CA, USA). The primer sequences used were designed to generate overlapping amplicons of approximately 1000 and 2000 nt in size and no modifications were made to the original CDV sequencing protocol. The end-repair and dA tailing were performed with the NEBNext Ultra II End Repair/dA-Tailing Module (New England Biolabs, USA). DNA barcodes EXP-NBD196, (Oxford Nanopore Technologies, UK) were ligated with NEBNext Ultra II Ligation Module (New England Biolabs, USA). After purification with Ampure XP beads, the AMII sequencing adapters were ligated with NEBNext Quick Ligation Module. Sixty ng of the final library was loaded onto a R9.4.1 (FLO-MIN106D) flow cell.
Also, consensus primers were designed to obtain the sequence of the 5′ (p2679 5′-ACCAGAMAAAGTTGGCTAWGGATAGW-3′) and 3′ genome (p2683R 5′-ACCAGACAAAGCTGGGTATGATAACT-3′) terminations.

2.2. Genome Assembly and Annotation

Base-calling and barcode demultiplexing were performed with Dorado version 1.1.0. Sequence reads below the expected size were removed and the consensus sequence was assembled by mapping to the MN267060 using the Geneious mapper (version Geneious Prime 2021.6.0.). Medaka (version 2022.1.1.) was used to map trimmed reads against a preliminary consensus to generate polished consensus sequences. The generated consensus sequences were manually checked for base-calling errors, especially in the homopolymeric regions. The genome was annotated using the ORF Finder (https://www.ncbi.nlm.nih.gov/orffinder/ (accessed on 5 September 2025)).

2.3. Phylogenetic Analysis

Sequences for the N, P, M, F, H, and L genes were first aligned using the MAFFT webserver with default parameters (https://www.ebi.ac.uk/jdispatcher/msa/mafft (accessed on 6 September 2025)). Thereafter, the IQ-TREE webserver (http://iqtree.cibiv.univie.ac.at/ (accessed on 6 September 2025)) was used to select substitution models and reconstruct maximum-likelihood phylogenetic trees, with 1000 ultrafast bootstrap replicates. The phylogenetic analyses were conducted under the GTR + F + I + G4 substitution model, which was identified as the best-fitting model according to the Akaike Information Criterion (AIC). Subsequently, the resulting trees were edited in the iTOL webserver (https://itol.embl.de/ (accessed on 7 September 2025)).

2.4. Recombination Detection

Whole-genome sequences were used to identify potential recombination events in Rockborn using the algorithms implemented in the Recombination Detection Program v.4.101 (RDP4) [24]. Default settings were used for each algorithm. A recombination event was accepted when detected by 7 distinct methods (RDP, GENECONV, BootScan, MaxChi, Chimaera, SiScan, 3Seq) implemented in the program, each with a p-value < 5 × 10⁻⁴.

2.5. Sequence Deposition

The genome sequence of the Rockborn strain was deposited in GenBank under the accession number PX254110.

3. Results

A total of 551.860 reads were mapped to CDV. The mean vertical coverage was about 44,000×. The assembled genome was 15,690 nt in length, with 107 and 105 nt long UTRs at the 3′ and 5′ ends, respectively. The lengths of N, P, M, F, H and L genes (and their respective proteins) were 1572 nt (523 aa), 1524 nt (507 aa), 1008 nt (335 aa), 1989 nt (662 aa), 1824 nt (607 aa) and 6555 nt (2184 aa), respectively. The genome and gene-wise sequence homologies showed values of over 98% and 99%, respectively, to the most closely related CDV strains, many, if not all, of which are putative Rockborn-derived vaccines and wild-type strains (Table 1).
Next, whole-genome and gene-wise phylogenies were carried out (Figure 1). A whole-genome-based alignment was assembled from 223 reference genome sequences available in the GenBank and the newly determined Rockborn sequence. After a preliminary analysis, 27 relevant sequences were retained for further processing; these sequences represented the main genetic clades. The genome-wise phylogeny indicated that strain Rockborn clusters with two field strains for which full-length genomes are available, the HN19 strain from a masked civet in China and the R252 strain from a dog in the United States. These features indicate that strain Rockborn, along with some recent field isolates, constitutes a unique genetic lineage alongside more deeply analyzed CDV lineages. A closer look revealed that R252 and HN19 CDVs share greater genetic relatedness across the entire genome, except for the region spanning the F and H genes, from nt positions 4935 to 6923 and 7079 to 8902, respectively, in the masked civet-origin CDV strain (GenBank ID: MT448054). In this data set, analysis of this genomic region showed that the masked civet-origin HN19 strain was more closely related to Rockborn than to the canine-origin R252 strain.
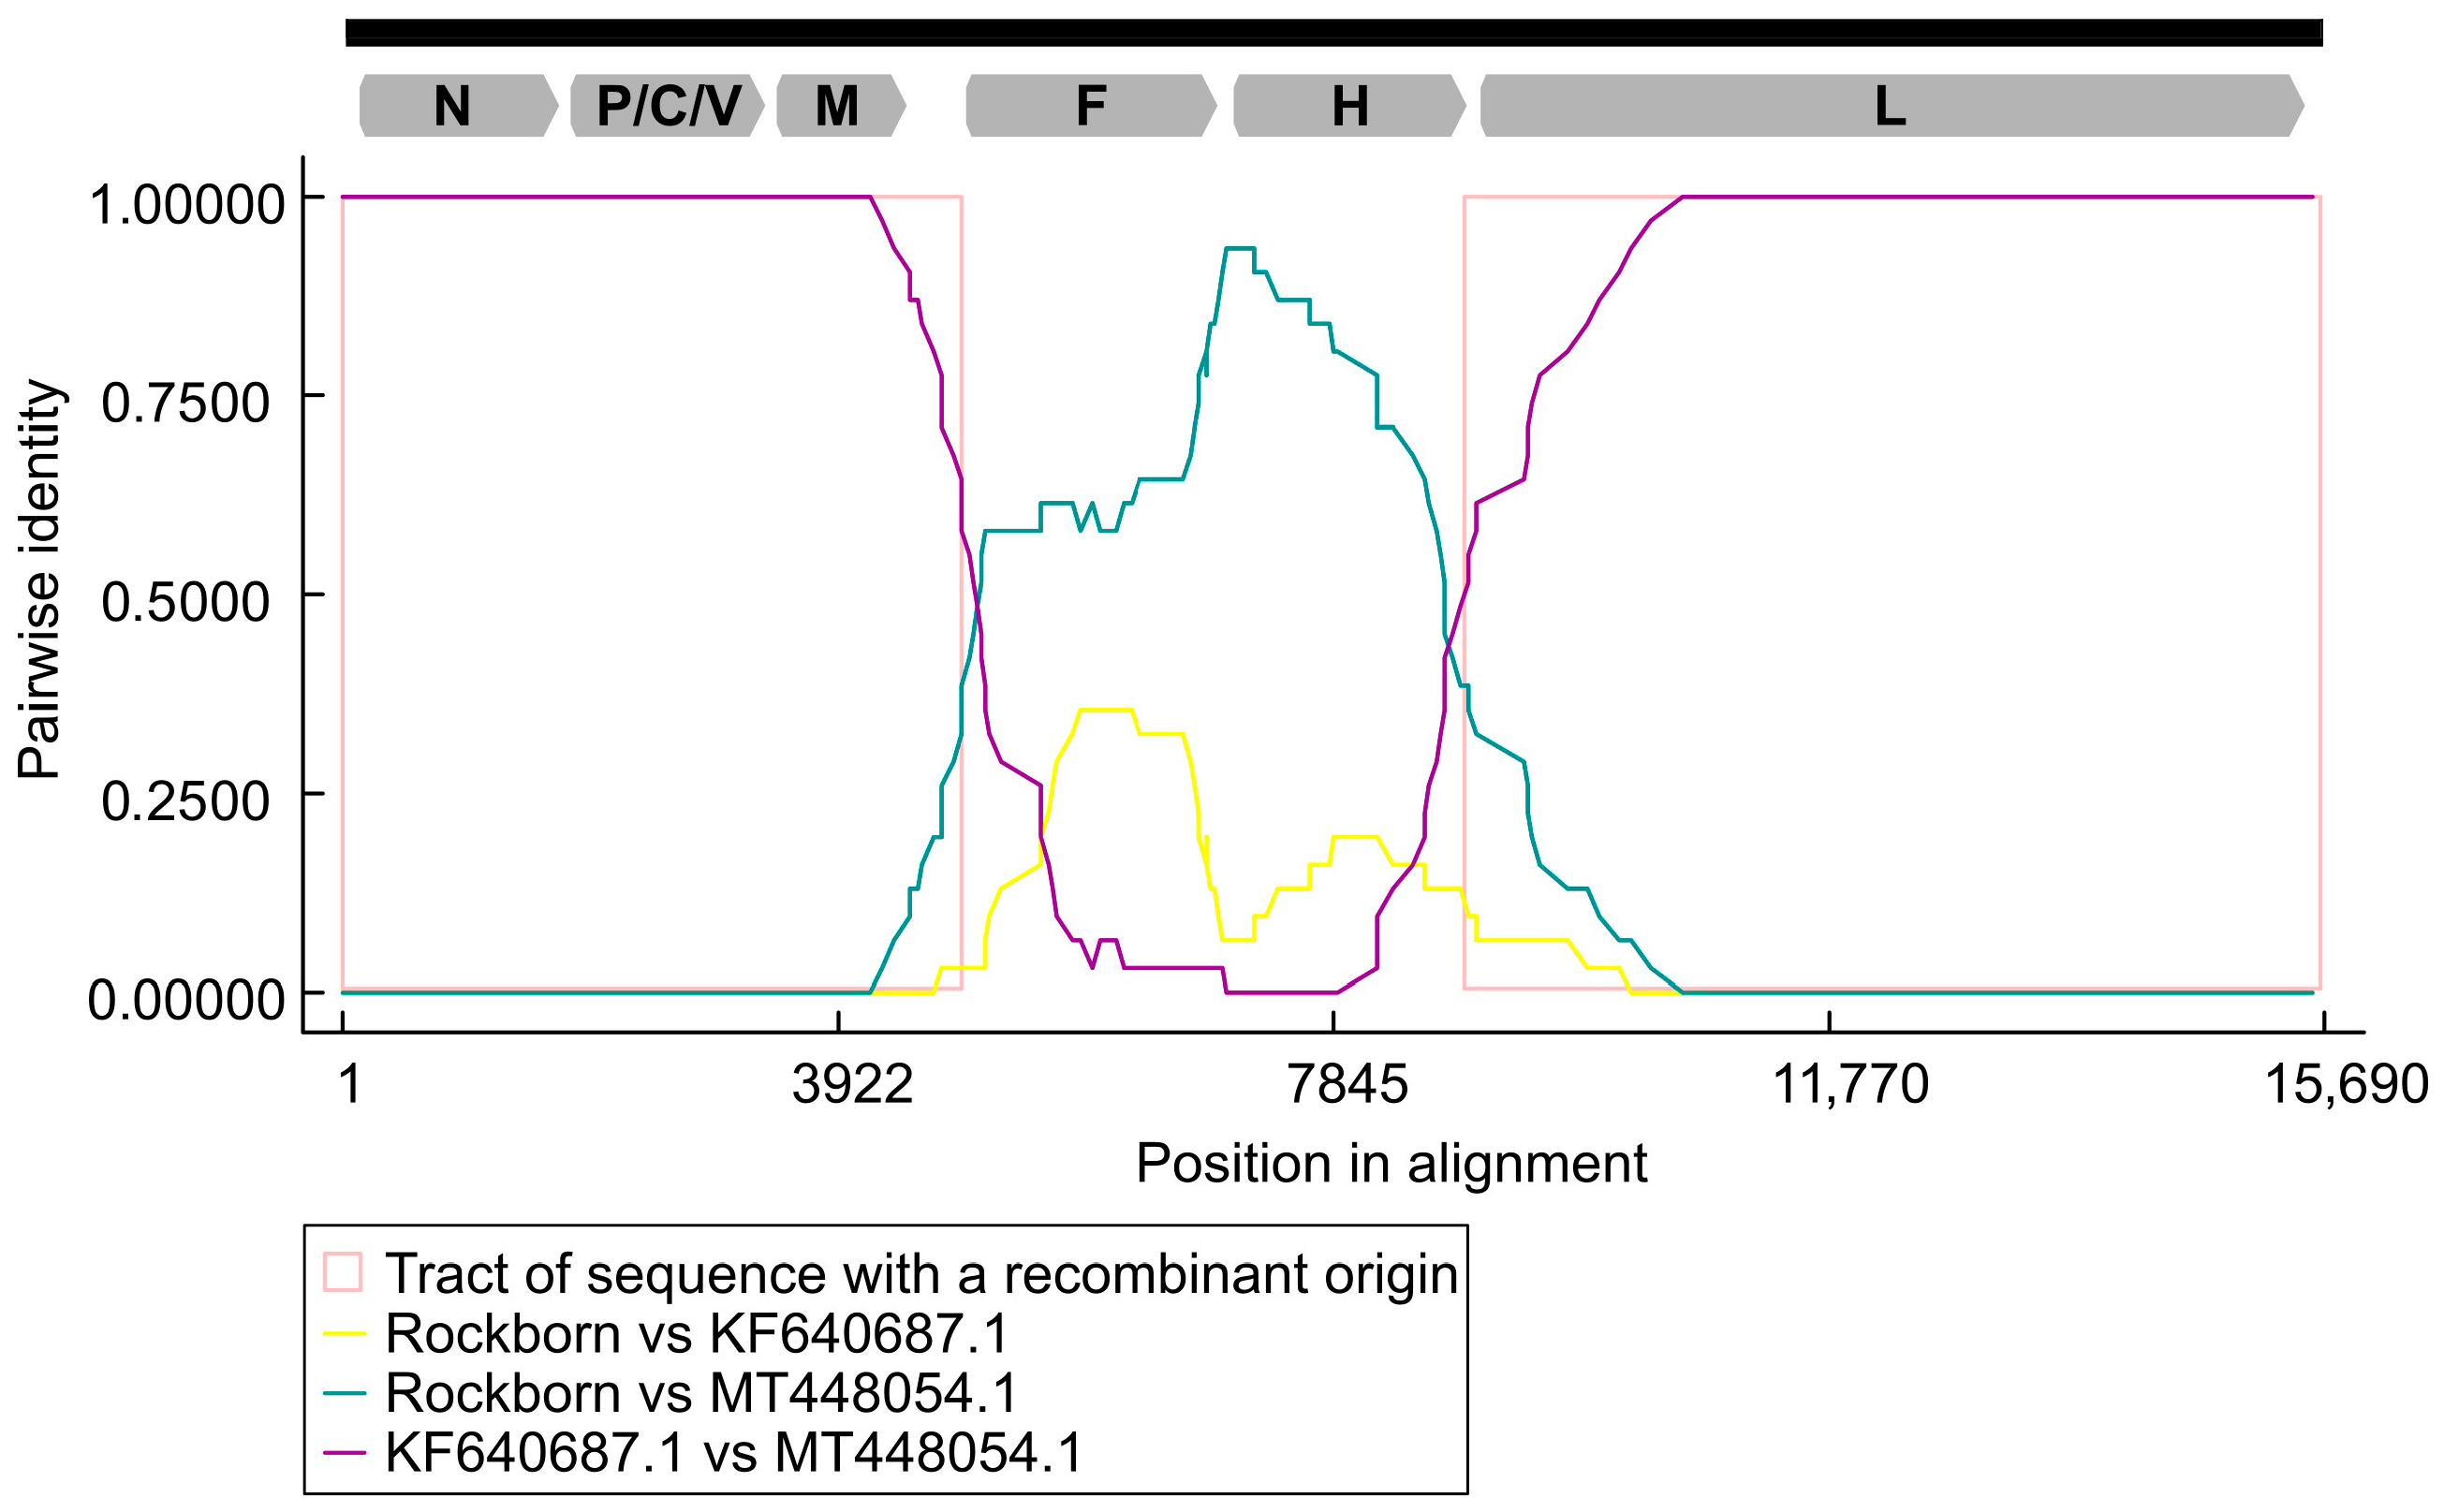
These findings suggest that past recombination events led to the mosaic structure of the genomes of some strains within the Rockborn lineage. This peculiar pattern of gene-wise relationship among Rockborn-like strains with available full-length genomes was confirmed by recombination analysis using dedicated bioinformatic analyses (Figure 2). We used the whole alignment containing 224 sequences as input to run the analysis with default parameters. As a result, an intra-lineage recombination event was observed within Rockborn-like strains (all but one algorithm showed evidence of recombination), placing the breakpoint sites at nt 4934 and 8902 (99% CI, 4747–5036 and 8482–8974, respectively). This region spans the entire F and H genes and may include short fragments of non-coding regions upstream and downstream, as indicated by the 99% CI values.

4. Discussion

This study presents the complete genome sequence of the historic CDV strain Rockborn, revealing its ongoing relevance in the contemporary epidemiological landscape. These new data not only resolve long-standing questions about the genetic makeup of this lineage beyond the H gene but also underscore the complex evolutionary forces, particularly recombination, that shape CDV diversity [17,21].
Given the official withdrawal of Rockborn-based vaccines from many markets in the mid-1990s due to safety concerns [19,20], a significant aspect of this whole-genome sequencing study is the confirmation that Rockborn-like viruses or Rockborn-derived recombinants continue to circulate in dogs and wildlife. Based on the H gene sequence, Martella and co-workers identified two vaccines on the market containing Rockborn-like strains and noted that several field isolates from Austria and Japan showed nearly 100% nucleotide identity to the Rockborn-46th laboratory strain [21]. More recent studies have continued to detect these strains, reinforcing the idea of their natural persistence within and among different animal populations. Based on whole-genome sequence data generated in our study and database interrogation, we obtained additional evidence for the presence of Rockborn-like CDV strains in dogs and other carnivores (Table 1). Rockborn-like CDV was identified in a captive vaccinated fennec fox in Japan, 2017 [25], in non-vaccinated farmed masked civets in China, 2019 [26]. Also, Rockborn-like CDV was reported repeatedly, although sporadically, in domestic dogs in Brazil, 2015 [27], Canada, 2025 [28], and New Zealand, 2021–2024 [29], suggesting a continual spill-over of vaccine-derived Rockborn-like CDV strains into the environment.
The central finding of our genomic analysis was the identification of a clear recombination event, positioning the Rockborn strain as a probable parental strain to a mosaic virus circulating in the wild. While recombination in negative-sense RNA viruses was once considered a rare phenomenon [30], a growing body of evidence now establishes it as a key force driving the evolution and genetic diversity of CDV [17,31,32]. Previous studies have extensively documented recombination events involving various CDV genotypes. Budaszewski and co-workers identified eight putative recombinant viruses, demonstrating that homologous recombination is a frequent event in natural CDV populations and that vaccine strains, particularly from the America-1 lineage, play a significant role in shaping viral evolution [32]. Similarly, Yuan et al. [31] concluded that recombination is a key evolutionary force after identifying six distinct events and noting that viruses isolated from different host species—such as minks, seals, and raccoons—can recombine, contributing to the broad host range and adaptability of CDVs. Our study demonstrates that the historic Rockborn lineage is actively participating in this process, dispersing genome fragments of the original strain into other CDV strains over the long term via recombination. The mosaic genome structure observed, with parental contributions from strains infecting distantly related hosts like masked civets and domestic dogs, exemplifies the cross-species nature of these evolutionary events. In this case, the masked civet strain HN19 was closely related to the canine strain R252 across the genome, but the F and H genes, which were likely derived from strain Rockborn. These genes encode key proteins involved in cell-to-cell viral spread (F protein) and receptor interactions (H protein) [7,8,9,10,11]. Understanding the effects of the exchange of cognate genome regions is challenging and requires multiple studies using reverse genetics, in vitro or in vivo experiments, or both, for functional validation. Also, sampling bias and the heavy reliance on publicly available GenBank sequences might affect our analysis or the findings of analogous studies.
This work strongly emphasizes the limitations of relying on single-gene analyses for CDV genotyping and diagnostics. The H gene has traditionally been a target of phylogenetic studies due to its high variability and its role in immunity. Consequently, many diagnostic assays are designed to differentiate wild-type and vaccine strains based on specific polymorphisms within the H gene, or alternatively, other genomic regions [16,33,34,35,36,37,38]. However, the occurrence of recombination means that a virus can possess a gene from one lineage/sub-lineage while other parts of its genome originate from another lineage/sub-lineage. For example, the recombination event identified in this study appeared to affect the F and H genes, suggesting that any diagnostic or phylogenetic analysis based solely on the H gene would have masked and mischaracterized the recombinant virus. This risk has been pointed out in other whole-genome studies. Budaszewski and co-workers emphasized that genotyping based on partial genomic sequences could lead to misidentification and that complete genome analysis is necessary to detect recombination hot spots, which they frequently found in the F and H genes [32]. Likewise, Yuan and co-workers identified recombination events primarily in the P and L genes, regions often overlooked in standard genotyping [31]. Therefore, relying on single-gene assays in a landscape shaped by recombination can lead to flawed epidemiological conclusions and ineffective control strategies.
Vaccine-related disease has been reported on several occasions in dogs, but sporadically. In most cases, however, it has been difficult to establish whether the disease was induced by the vaccine virus or by field viruses infecting pups shortly before or after vaccine administration. This issue could not be ruled out definitively in earlier studies, as appropriate diagnostic tools were not available. Using a genotyping PCR assay that distinguishes between locally relevant CDV genotypes and vaccine viruses, all suspected cases of CDV vaccine-induced illness or death analyzed during a 2-year surveillance in Italy were caused by CDV field strains [16], suggesting that vaccine-induced residual virulence is rare. Optimized diagnostic tools capable of distinguishing between field and vaccine CDV strains could help better assess the magnitude of this phenomenon.

5. Conclusions

In conclusion, the full-genome characterization of the Rockborn strain not only fills a critical gap in our understanding of this historic lineage but also reveals its ongoing role in shaping the complex evolutionary dynamics of CDV through recombination. This underscores the need to shift towards whole-genome approaches for effective surveillance, accurate diagnostics, and a comprehensive understanding of CDV epidemiology and evolution.

Supplementary Materials

The following supporting information can be downloaded at: https://www.mdpi.com/article/10.3390/vetsci13010081/s1, Table S1: Functions of canine distemper virus genes.

Author Contributions

Conceptualization, V.M. and N.D.; methodology, Z.L., K.B., Á.B., G.D., G.L. and F.P.; validation, G.K. and V.M.; formal analysis, Z.L., K.B., Á.B., G.D., G.L. and F.P.; resources, G.K. and V.M.; data curation, Z.L., G.L. and V.M.; writing—original draft preparation, Z.L., K.B. and V.M.; writing—review and editing, Z.L., K.B., Á.B., G.D., G.L., G.K., F.P. and V.M.; visualization, Z.L.; supervision, G.K. and V.M.; project administration, G.L. and N.D.; funding acquisition, Z.L., K.B., G.K. and V.M. All authors have read and agreed to the published version of the manuscript.

Funding

V.M. and K.B. were supported by the National Research, Development and Innovation Office, Hungary under grant RRF-2.3.1-21-2022-00001 (National Laboratory for Infectious Animal Diseases, Antimicrobial Resistance, Veterinary Public Health and Food Chain Safety). Z.L., G.K., and the mobilization of sequencing tools were supported by Project no. TKP2021-NVA-07, which has been implemented with the support provided from the National Research, Development and Innovation Fund of Hungary, financed under the TKP2021-NVA funding scheme. Z.L. was a recipient of the János Bolyai Research Scholarship of the Hungarian Academy of Sciences (BO/00124/24).

Institutional Review Board Statement

Not applicable.

Informed Consent Statement

Not applicable.

Data Availability Statement

The data presented in this study are openly available. The genome sequence of the Rockborn strain was deposited in the Genbank (accession number, PX254110).

Acknowledgments

The authors have reviewed and edited the output and take full responsibility for the content of this publication.

Conflicts of Interest

The authors declare no conflicts of interest.

References

  1. Sourimant, J.; Plemper, R.K. Organization, Function, and Therapeutic Targeting of the Morbillivirus RNA-Dependent RNA Polymerase Complex. Viruses 2016, 8, 251. [Google Scholar] [CrossRef] [Scilit]
  2. Barrett, T.; Shrimpton, S.B.; Russell, S.E.H. Nucleotide Sequence of the Entire Protein Coding Region of Canine Distemper Virus Polymerase-Associated (P) Protein MRNA. Virus Res. 1985, 3, 367–372. [Google Scholar] [CrossRef] [Scilit]
  3. Röthlisberger, A.; Wiener, D.; Schweizer, M.; Peterhans, E.; Zurbriggen, A.; Plattet, P. Two Domains of the V Protein of Virulent Canine Distemper Virus Selectively Inhibit STAT1 and STAT2 Nuclear Import. J. Virol. 2010, 84, 6328–6343. [Google Scholar] [CrossRef] [Scilit]
  4. Siering, O.; Sawatsky, B.; Pfaller, C.K. C Protein Is Essential for Canine Distemper Virus Virulence and Pathogenicity in Ferrets. J. Virol. 2021, 95, 4. [Google Scholar] [CrossRef] [Scilit]
  5. Dietzel, E.; Anderson, D.E.; Castan, A.; von Messling, V.; Maisner, A. Canine Distemper Virus Matrix Protein Influences Particle Infectivity, Particle Composition, and Envelope Distribution in Polarized Epithelial Cells and Modulates Virulence. J. Virol. 2011, 85, 7162–7168. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  6. Anderson, D.E.; von Messling, V. Region between the Canine Distemper Virus M and F Genes Modulates Virulence by Controlling Fusion Protein Expression. J. Virol. 2008, 82, 10510–10518. [Google Scholar] [CrossRef] [Scilit]
  7. von Messling, V.; Cattaneo, R. Amino-Terminal Precursor Sequence Modulates Canine Distemper Virus Fusion Protein Function. J. Virol. 2002, 76, 4172–4180. [Google Scholar] [CrossRef] [Scilit]
  8. Plattet, P.; Cherpillod, P.; Wiener, D.; Zipperle, L.; Vandevelde, M.; Wittek, R.; Zurbriggen, A. Signal Peptide and Helical Bundle Domains of Virulent Canine Distemper Virus Fusion Protein Restrict Fusogenicity. J. Virol. 2007, 81, 11413–11425. [Google Scholar] [CrossRef] [Scilit]
  9. Pratakpiriya, W.; Seki, F.; Otsuki, N.; Sakai, K.; Fukuhara, H.; Katamoto, H.; Hirai, T.; Maenaka, K.; Techangamsuwan, S.; Lan, N.T.; et al. Nectin4 Is an Epithelial Cell Receptor for Canine Distemper Virus and Involved in Neurovirulence. J. Virol. 2012, 86, 10207–10210. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  10. Alves, L.; Khosravi, M.; Avila, M.; Ader-Ebert, N.; Bringolf, F.; Zurbriggen, A.; Vandevelde, M.; Plattet, P. SLAM- and Nectin-4-Independent Noncytolytic Spread of Canine Distemper Virus in Astrocytes. J. Virol. 2015, 89, 5724–5733. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  11. Zhao, J.; Ren, Y. Multiple Receptors Involved in Invasion and Neuropathogenicity of Canine Distemper Virus: A Review. Viruses 2022, 14, 1520. [Google Scholar] [CrossRef] [Scilit]
  12. MacLachlan, N.; Dubovi, E.; Fenner, F. Paramyxoviridae. In Fenner’s Veterinary Virology; Academic Press: Boston, MA, USA, 2011; pp. 299–325. [Google Scholar]
  13. Rima, B.; Balkema-Buschmann, A.; Dundon, W.G.; Duprex, P.; Easton, A.; Fouchier, R.; Kurath, G.; Lamb, R.; Lee, B.; Rota, P.; et al. ICTV Virus Taxonomy Profile: Paramyxoviridae. J. Gen. Virol. 2019, 100, 1593–1594. [Google Scholar] [CrossRef] [Scilit]
  14. Blixenkrone-Möller, M.; Svansson, V.; Appel, M.; Krogsrud, J.; Have, P.; Örvell, C. Antigenic Relationships between Field Isolates of Morbilliviruses from Different Carnivores. Arch. Virol. 1992, 123, 279–294. [Google Scholar] [CrossRef] [Scilit]
  15. Iwatsuki, K.; Tokiyoshi, S.; Hirayama, N.; Nakamura, K.; Ohashi, K.; Wakasa, C.; Mikami, T.; Kai, C. Antigenic Differences in the H Proteins of Canine Distemper Viruses. Vet. Microbiol. 2000, 71, 281–286. [Google Scholar] [CrossRef] [Scilit]
  16. Martella, V.; Elia, G.; Lucente, M.; Decaro, N.; Lorusso, E.; Banyai, K.; Blixenkronemoller, M.; Lan, N.; Yamaguchi, R.; Cirone, F. Genotyping Canine Distemper Virus (CDV) by a Hemi-Nested Multiplex PCR Provides a Rapid Approach for Investigation of CDV Outbreaks. Vet. Microbiol. 2007, 122, 32–42. [Google Scholar] [CrossRef] [Scilit]
  17. Ke, G.-M.; Ho, C.-H.; Chiang, M.-J.; Sanno-Duanda, B.; Chung, C.-S.; Lin, M.-Y.; Shi, Y.-Y.; Yang, M.-H.; Tyan, Y.-C.; Liao, P.-C.; et al. Phylodynamic Analysis of the Canine Distemper Virus Hemagglutinin Gene. BMC Vet. Res. 2015, 11, 164. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  18. Rockborn, G. Canine Distemper Virus in Tissue Culture. Arch. Gesamte Virusforsch. 1958, 8, 485–492. [Google Scholar] [CrossRef] [Scilit]
  19. Cornwell, H.; Thompson, H.; McCandlish, I.; Macartney, L.; Nash, A. Encephalitis in Dogs Associated with a Batch of Canine Distemper (Rockborn) Vaccine. Vet. Rec. 1988, 122, 54–59. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  20. Gloyd, J. Vaccines Recalled. J. Am. Vet. Med. Assoc. 1995, 207, 1397. [Google Scholar]
  21. Martella, V.; Blixenkrone-Møller, M.; Elia, G.; Lucente, M.S.; Cirone, F.; Decaro, N.; Nielsen, L.; Bányai, K.; Carmichael, L.E.; Buonavoglia, C. Lights and Shades on an Historical Vaccine Canine Distemper Virus, the Rockborn Strain. Vaccine 2011, 29, 1222–1227. [Google Scholar] [CrossRef] [Scilit]
  22. Appel, M.J.G. Reversion to Virulence of Attenuated Canine Distemper Virus In Vivo and In Vitro. J. Gen. Virol. 1978, 41, 385–393. [Google Scholar] [CrossRef] [Scilit]
  23. Lanszki, Z.; Tóth, G.E.; Schütz, É.; Zeghbib, S.; Rusvai, M.; Jakab, F.; Kemenesi, G. Complete Genomic Sequencing of Canine Distemper Virus with Nanopore Technology during an Epizootic Event. Sci. Rep. 2022, 12, 4116. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  24. Martin, D.P.; Murrell, B.; Golden, M.; Khoosal, A.; Muhire, B. RDP4: Detection and Analysis of Recombination Patterns in Virus Genomes. Virus Evol. 2015, 1, vev003. [Google Scholar] [CrossRef] [Scilit]
  25. Tamukai, K.; Minami, S.; Kurihara, R.; Shimoda, H.; Mitsui, I.; Maeda, K.; Une, Y. Molecular Evidence for Vaccine-Induced Canine Distemper Virus and Canine Adenovirus 2 Coinfection in a Fennec Fox. J. Vet. Diagn. Investig. 2020, 32, 598–603. [Google Scholar] [CrossRef] [Scilit]
  26. Shi, N.; Zhang, L.; Yu, X.; Zhu, X.; Zhang, S.; Zhang, D.; Duan, M. Insight Into an Outbreak of Canine Distemper Virus Infection in Masked Palm Civets in China. Front. Vet. Sci. 2021, 8, 728238. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  27. Freitas, L.A.; Leme, R.A.; Saporiti, V.; Alfieri, A.A.; Alfieri, A.F. Molecular Analysis of the Full-Length F Gene of Brazilian Strains of Canine Distemper Virus Shows Lineage Co-Circulation and Variability between Field and Vaccine Strains. Virus Res. 2019, 264, 8–15. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  28. Rätsep, E.; Ojkic, D. Canine Distemper Virus Infection of Vaccinal Origin in a 14-Week-Old Puppy. J. Vet. Diagn. Investig. 2024, 36, 287–290. [Google Scholar] [CrossRef] [Scilit]
  29. Gulliver, E.; Taylor, H.; Eames, M.; Chernyavtseva, A.; Jauregui, R.; Wilson, A.; Bestbier, M.; O’Connell, J.; Buckle, K.; Castillo-Alcala, F. Investigation of Post-Vaccinal Canine Distemper Involving the Rockborn-like Strain in Nine Puppies in New Zealand. N. Z. Vet. J. 2025, 73, 278–287. [Google Scholar] [CrossRef] [Scilit]
  30. Chare, E.R.; Gould, E.A.; Holmes, E.C. Phylogenetic Analysis Reveals a Low Rate of Homologous Recombination in Negative-Sense RNA Viruses. J. Gen. Virol. 2003, 84, 2691–2703. [Google Scholar] [CrossRef] [Scilit]
  31. Yuan, C.; Liu, W.; Wang, Y.; Hou, J.; Zhang, L.; Wang, G. Homologous Recombination Is a Force in the Evolution of Canine Distemper Virus. PLoS ONE 2017, 12, e0175416. [Google Scholar] [CrossRef] [Scilit]
  32. da Fontoura Budaszewski, R.; Streck, A.F.; Nunes Weber, M.; Maboni Siqueira, F.; Muniz Guedes, R.L.; Wageck Canal, C. Influence of Vaccine Strains on the Evolution of Canine Distemper Virus. Infect. Genet. Evol. 2016, 41, 262–269. [Google Scholar] [CrossRef] [Scilit]
  33. Sui, P.; Sun, Y.; Shi, Y.; Ran, W.; Shi, N.; Sun, D.; Zheng, J.; Zhao, J. Establishment and Evaluation of a Multiplex Real-Time RT-PCR for Quantitative and Differential Detection of Wild-Type Canine Distemper Virus from Vaccine Strains. Heliyon 2023, 9, e19344. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  34. Dong, X.Y.; Li, W.H.; Zhu, J.L.; Liu, W.J.; Zhao, M.Q.; Luo, Y.W.; Chen, J.D. Detection and Differentiation of Wild-Type and Vaccine Strains of Canine Distemper Virus by a Duplex Reverse Transcription Polymerase Chain Reaction. Iran. J. Vet. Res. 2015, 16, 172–175. [Google Scholar] [PubMed]
  35. Si, W.; Zhou, S.; Wang, Z.; Cui, S. A Multiplex Reverse Transcription-Nested Polymerase Chain Reaction for Detection and Differentiation of Wild-Type and Vaccine Strains of Canine Distemper Virus. Virol. J. 2010, 7, 86. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  36. Yi, L.; Cheng, S.; Xu, H.; Wang, J.; Cheng, Y.; Yang, S.; Luo, B. Development of a Combined Canine Distemper Virus Specific RT-PCR Protocol for the Differentiation of Infected and Vaccinated Animals (DIVA) and Genetic Characterization of the Hemagglutinin Gene of Seven Chinese Strains Demonstrated in Dogs. J. Virol. Methods 2012, 179, 281–287. [Google Scholar] [CrossRef] [Scilit]
  37. Wilkes, R.P.; Sanchez, E.; Riley, M.C.; Kennedy, M.A. Real-Time Reverse Transcription Polymerase Chain Reaction Method for Detection of Canine Distemper Virus Modified Live Vaccine Shedding for Differentiation from Infection with Wild-Type Strains. J. Vet. Diagn. Investig. 2014, 26, 27–34. [Google Scholar] [CrossRef] [Scilit]
  38. Liu, D.-F.; Liu, C.-G.; Tian, J.; Jiang, Y.-T.; Zhang, X.-Z.; Chai, H.-L.; Yang, T.-K.; Yin, X.-C.; Zhang, H.-Y.; Liu, M.; et al. Establishment of Reverse Transcription Loop-Mediated Isothermal Amplification for Rapid Detection and Differentiation of Canine Distemper Virus Infected and Vaccinated Animals. Infect. Genet. Evol. 2015, 32, 102–106. [Google Scholar] [CrossRef] [Scilit]
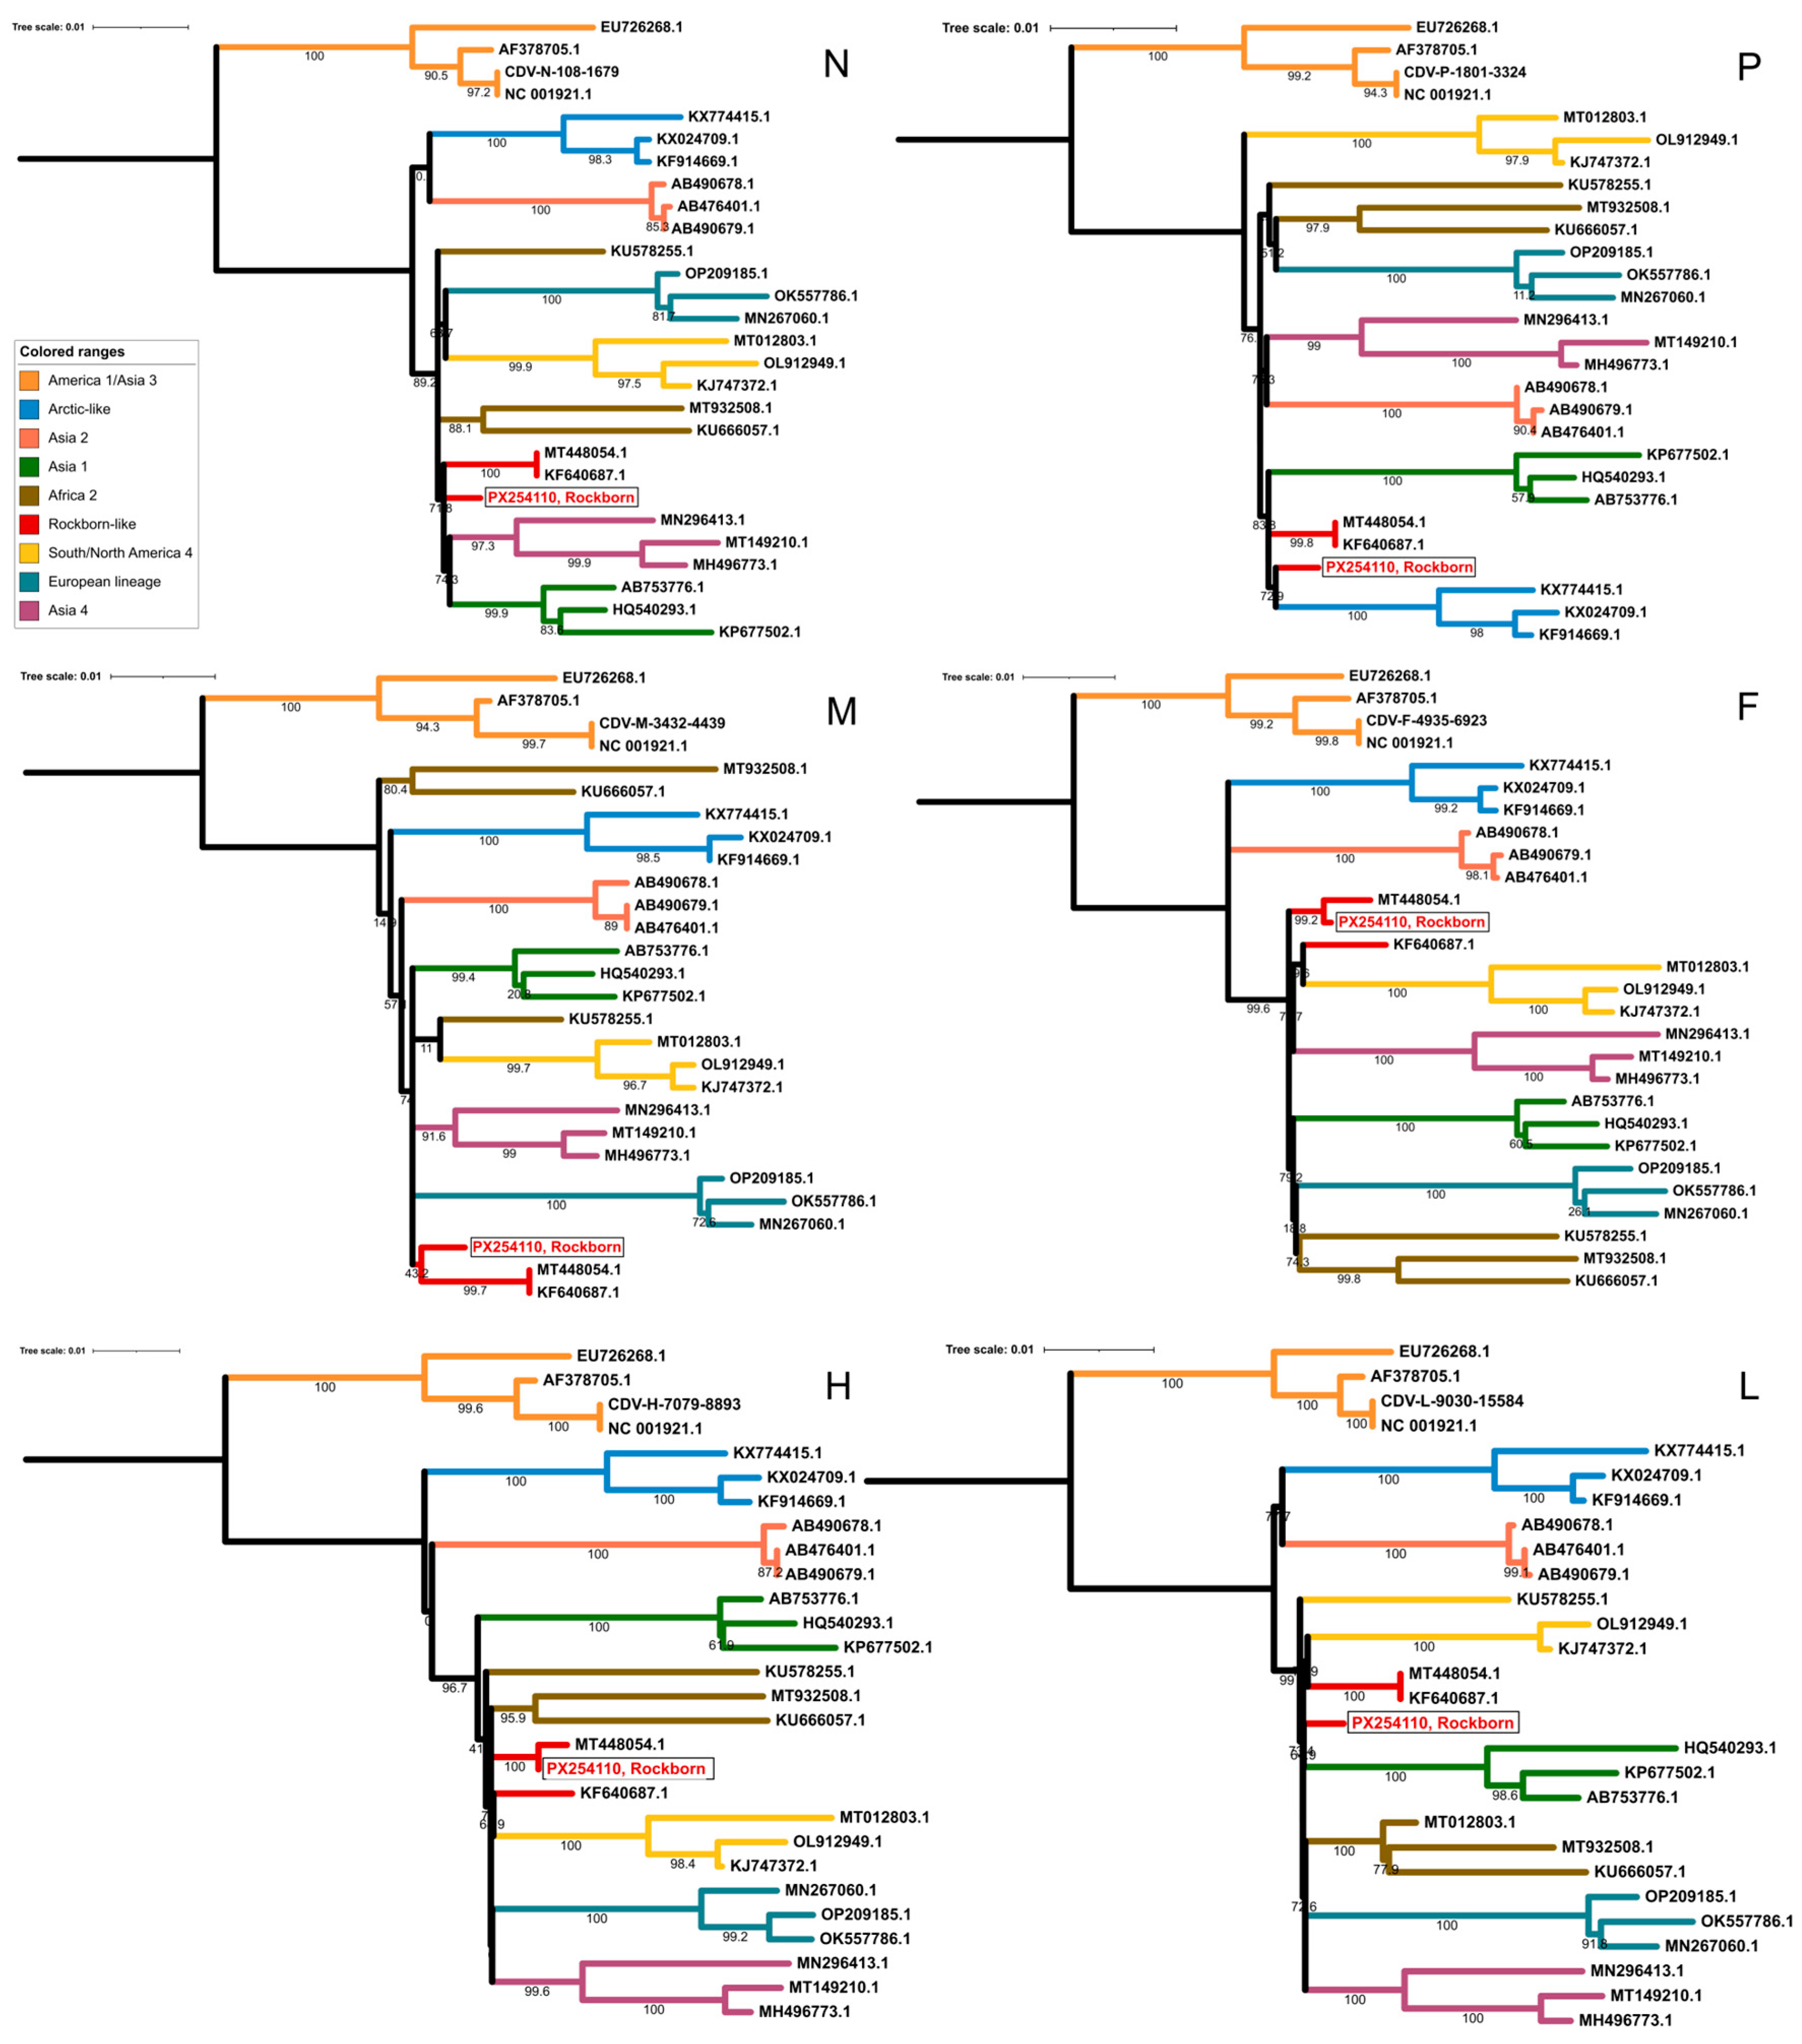
Figure 1. Gene-wise maximum-likelihood phylogenies based on 28 CDV sequences for the N, P, M, F, H, and L genes. Major CDV lineages are color-coded, and the Rockborn strain is highlighted in red and boxed. The clustering patterns of the Rockborn strain in the F and H genes are indicative of a recombination event. Bootstrap support values are shown at key nodes.
Vetsci 13 00081 g001
Figure 2. Corrected pairwise identity analysis generated using the RDP4 software (version 4.4). Pairwise identity plot obtained by comparing the Rockborn-46th strain with two closely related CDV genomes, HN19 (MT448054.1) and R252 (KF640687.1), showing identity values across the full genome alignment and the corresponding breakpoint confidence intervals. Different colors are used to depict the different identity plots. The genome structure is also shown.
Vetsci 13 00081 g002
Table 1. CDV strains with the greatest sequence homology in the GenBank to strain Rockborn. Strains with uncertain metadata are indicated by *.
GeneGenBank Entry, nt IdentityOrigin
Host SpeciesGeographicYear
Whole genomeMT448054, 98.8%Masked civetChina2019
KF640687, 98.5%DogUnited States1970s *
AF164967, 98.4%DogUnited States *1975 *
EU716337, 98.2%DogUnited States2004
NEU072200, 99.5%vaccine-2006
AY649446, 99.1%RaccoonUnited States2001
PEU072201, 99.7%vaccine-2006
KF640687, 99.1%DogUnited States1970s *
MT448054, 99.1%Masked civetChina2019
MEU072199, 99.7%vaccine-2006
FKY057355, 99.8%DogBrazil2015
EU072198, 99.3%vaccine-2006
MT448054, 99.3%Masked civetChina2019
HGU810819, 99.9%Rockborn, 46th cell passage-2011
LC498611, 99.8%Fennec foxJapan2017
GU266280, 99.8%vaccine-2011
FJ705238, 99.8%vaccine-2010
EF095750, 99.8%vaccine-2010
FJ461702, 99.7%vaccine-2008
MT448054, 99.6%Masked civetChina2019
AF178039, 99.5%Lesser pandaChina1999
OQ282897, 99.8%DogChina2021
OQ282900, 99.7%DogChina2021
OQ282902, 99.6%DogChina2021
JX912968, 99.3%DogBrazil2008
Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content.