Review

Pompe Disease: New Developments in an Old Lysosomal Storage Disorder

Cell and Developmental Biology Center, National Heart, Lung, and Blood Institute, NIH, Bethesda, MD 20892, USA
*
Author to whom correspondence should be addressed.
Biomolecules 2020, 10(9), 1339; https://doi.org/10.3390/biom10091339
Submission received: 27 August 2020 / Revised: 14 September 2020 / Accepted: 15 September 2020 / Published: 18 September 2020

Abstract

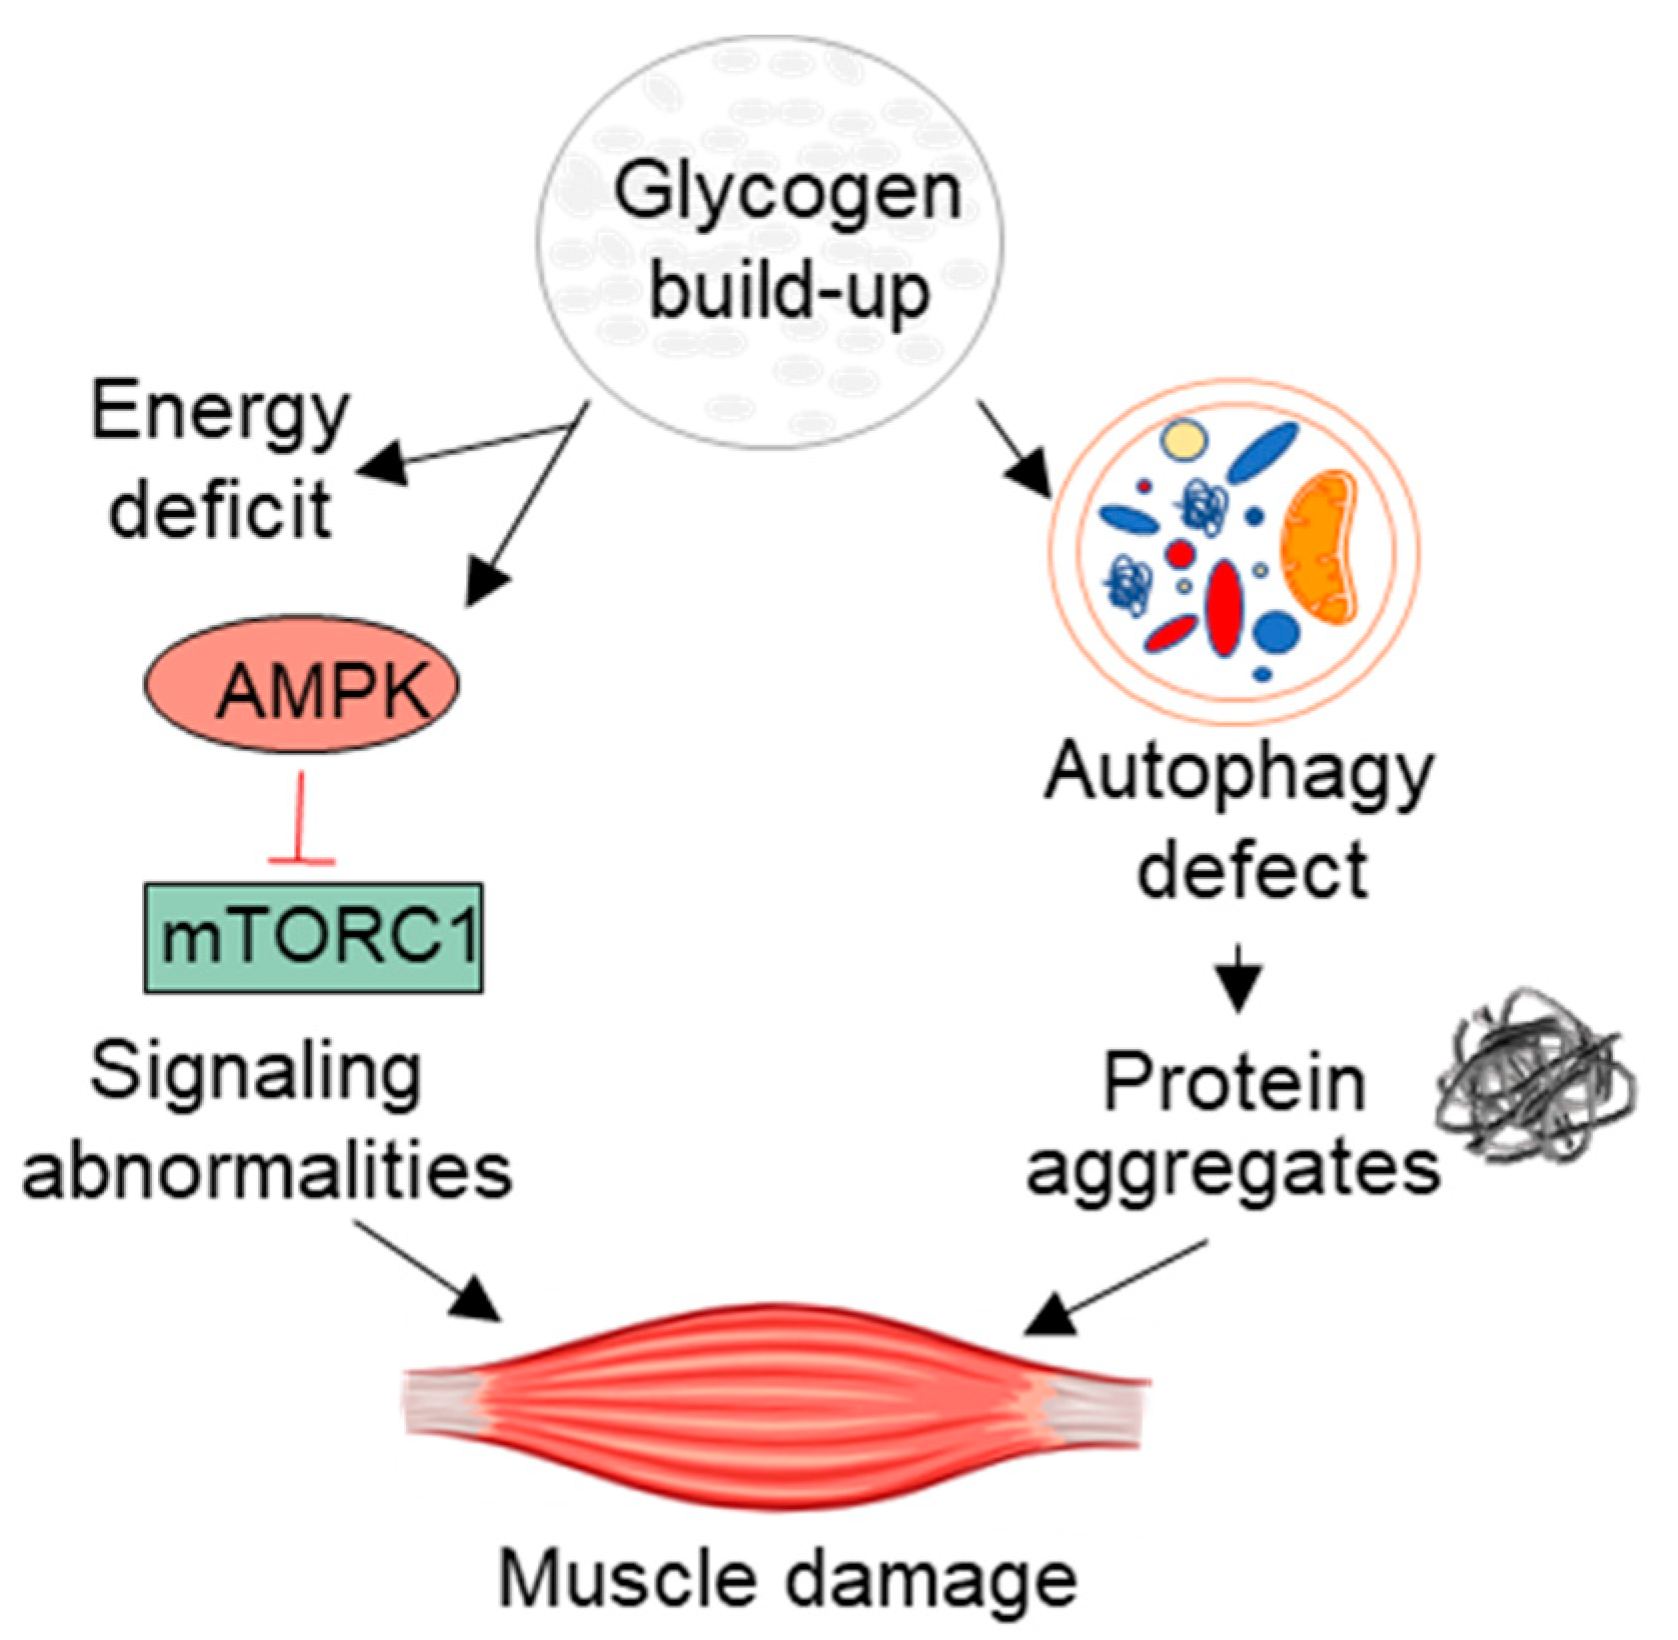
Pompe disease, also known as glycogen storage disease type II, is caused by the lack or deficiency of a single enzyme, lysosomal acid alpha-glucosidase, leading to severe cardiac and skeletal muscle myopathy due to progressive accumulation of glycogen. The discovery that acid alpha-glucosidase resides in the lysosome gave rise to the concept of lysosomal storage diseases, and Pompe disease became the first among many monogenic diseases caused by loss of lysosomal enzyme activities. The only disease-specific treatment available for Pompe disease patients is enzyme replacement therapy (ERT) which aims to halt the natural course of the illness. Both the success and limitations of ERT provided novel insights in the pathophysiology of the disease and motivated the scientific community to develop the next generation of therapies that have already progressed to the clinic.
Keywords: Pompe disease; lysosome; lysosomal targeting; autophagy; enzyme replacement therapy; gene therapy; muscle; satellite cells Pompe disease; lysosome; lysosomal targeting; autophagy; enzyme replacement therapy; gene therapy; muscle; satellite cells