1. Introduction
Unlike in chloride systems, Cu(II) is generally not considered an oxidant in the leaching of chalcopyrite in sulfate systems. This is due to the lack of stability of Cu(I) in sulfate media. However, some studies [
1,
2,
3,
4] observed that Cu(II) had an enhancing effect on chalcopyrite dissolution in sulfate media. Hiroyoshi et al. [
2] proposed a two-step process in which chalcopyrite is first reduced to chalcocite by Fe(II) in the presence of Cu(II) (Equation (1)). Subsequently, chalcocite is oxidized by Fe(III) to Cu(II) and S (Equation (2)). Notably, Hiroyoshi et al. [
2] observed the enhancing effect of Cu(II) only in presence of both Fe(II) and Fe(III). However, they did not show direct evidence of chalcocite formation [
1,
2].
The mechanism by Hiroyoshi et al. [
2] was refuted by Nicol and Lázaro [
5] who proposed a different, two-step mechanism in which the first step is the non-oxidative dissolution of chalcopyrite (Equation (3)), followed by the oxidation of H
2S by Fe(III) (Equation (4)). Nicol and Lázaro [
5] referred to this mechanism as a “non-oxidative/oxidative” reaction mechanism. According to these authors, this non-oxidative/oxidative mechanism could occur in parallel with the direct oxidative dissolution (Equation (5)).
While both mechanisms could explain chalcopyrite dissolution at low potentials, Nicol and Lázaro [
5] argue that, based on thermodynamic principles, the first step of the mechanism proposed by Hiroyoshi et al. [
2] (Equation (1)) was unlikely. Also, Nicol et al. [
6] pointed out that in the model suggested by Hiroyoshi and co-workers [
2], chalcopyrite reduction (Equation (1)) was proposed at potentials in which the process is unlikely to occur from both a kinetic and thermodynamic standpoint. Lu et al. [
7] agreed to the possibility of the formation of H
2S and CuS intermediates. Although they calculated a very low K value, they rationalized it by stating that this was feasible at low concentrations of H
2S.
The non-oxidative/oxidative mechanism [
6] was further extended to chloride media by Nicol et al. The authors observed that Cu(II) catalyzed the oxidation of H
2S in the chloride system. They postulated the formation of covellite by non-oxidative acidic attack of chalcopyrite (Equation (6)) as the first step of the overall dissolution of chalcopyrite in chloride media at potentials below 600 mV. H
2S formed during this step can form a Cu(II)–HS complex (Equation (7)), which is subject to further oxidation by oxygen (Equation (8)). The authors argued that the oxidation of this complex by oxygen would be faster than the direct oxidation of H
2S (Equation (9)).
Thus, the study of H
2S oxidation in acidic media is of interest in the leaching of chalcopyrite, especially when considered in the context of its applicability in heap leaching. It is worth highlighting that although Nicol and Lázaro [
5] pointed out that the oxidation of H
2S is a necessary subsequent step for the non-oxidative dissolution process to occur, the authors did not speak directly to the effect of Cu(II) in the process. In a sulfate system, it would be important to understand better the H
2S oxidation by Fe(III), which is the most common oxidant. Likewise, Cu(II) was shown to catalyze H
2S oxidation by O
2 in chloride media, but this catalytic effect of Cu(II) has still to be observed in sulfate media, particularly with Fe(III) as the oxidant.
Although not always studied in the context of chalcopyrite leaching, the oxidation of dissolved hydrogen sulfide (H
2S) by Fe(III) according to the reaction (4) is fairly well known. The addition of Fe(III) salt is used as a method to control sulfide in wastewater [
8]. Sulfur precipitates as S (Equation (4)), and Fe(II) readily reacts with either H
2S or HS
− (formed through H
2S dissociation, Equation (10), pka = 6.9) to form pyrrhotite (FeS) (Equations (11) and (12)) or other Fe(II) sulfides such as pyrite and marcasite (FeS
2) [
8,
9].
In this work, the role of Cu(II) in the leaching of chalcopyrite in sulfate media was studied through electrochemical methods and controlled leach tests aimed at investigating some aspects of the non-oxidative leaching of chalcopyrite and the oxidation of H2S. This investigation aimed to obtain a better understanding of the overall dissolution of chalcopyrite, particularly through a pathway that may involve the formation and further reaction of H2S in acidic sulfate media which do not contain complexing agents of Cu(II) and Cu(I).
3. Results
3.1. Formation of H2S During Non-Oxidative Leaching
The presence of H
2S in sulfate media containing Cu(II) during non-oxidative chalcopyrite leaching was tested by linear voltammetry, since the direct measurement of H
2S can be challenging owing to the very low expected concentrations.
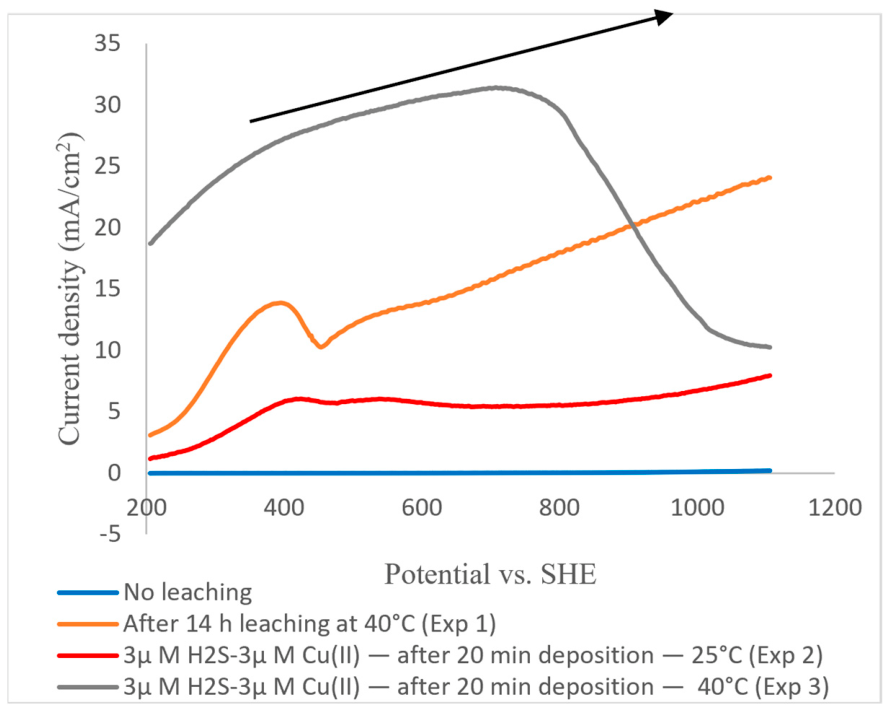
Figure 1 shows the results of the anodic stripping for a 0.1 M acid concentration. Anodic stripping commenced after the chalcopyrite electrode had been leached under nitrogen for 14 h at open circuit potential at 40 °C. A peak was observed around 420 mV (experiment 1). A similar peak but less intense was also observed for the solution initially containing 3µM of both Cu(II) and H
2S after prior deposition at 25 °C (experiment 2). This peak was accompanied by another at around 550 mV, both forming almost a single band. However, when the temperature for the deposition was increased to 40 °C (experiment 3), a single broad peak ranging from 200 to 1000 mV and with a maximum around 730mV was observed. It was more intense than the peak observed after the leaching of the chalcopyrite electrode at the same temperature. The similarity of the curves after the completion of experiments 1 and 2 suggests the presence of similar compounds in solution for both experiments. This indicates that both Cu(II) and H
2S were formed during the leaching of the chalcopyrite electrode at open circuit at 40 °C. The intensity of these peaks may be an indication of both Cu(II) and H
2S concentrations. This is in agreement with Nicol and Lazaro [
5], who used the same method, albeit with a more positive deposition potential and a different setup (ring-disk electrode), and came to the same conclusions. They attributed the peak they observed at 400 mV to the oxidation of Cu
2S formed by the reduction of Cu(II) in presence of H
2S (Equation (13)) on the platinum ring after open-circuit dissolution for the chalcopyrite disk.
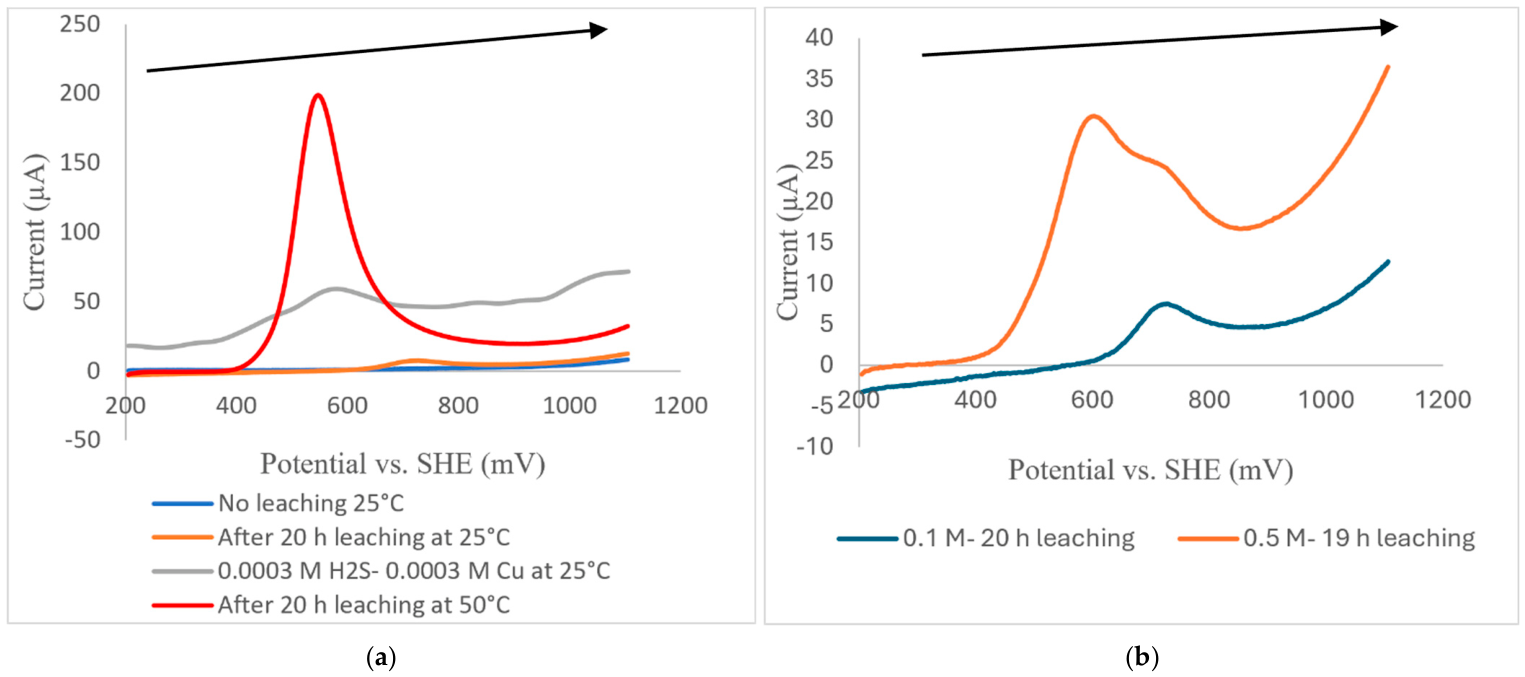
Similar results were observed with direct linear sweep voltammetry experiments without prior deposition (
Figure 2). For the acid concentration of 0.1 M (
Figure 2a), after 20 h of leaching at 25 °C, a small peak was observed around 700 mV. When the leaching temperature was raised to 50 °C, the peak became more intense and was shifted to the left, appearing at 550 mV. The peak appeared at almost the same position, around 600 mV, for the solution already containing both H
2S and Cu(II) but was less intense than after the non-oxidative leaching at 50 °C. The peak observed in all these conditions is likely due to the oxidation of a Cu–S compound containing Cu at +1 oxidation state and may be formed by reaction (Equation (13)). The comparison of current responses obtained after non-oxidative leaching at 0.1 and 0.5 M acid concentration (
Figure 2b) shows the peak was more intense and shifted to lower potentials at high acid strength. The difference of intensities suggests that the non-oxidative dissolution of chalcopyrite was promoted by high acid concentration. This is in agreement with the kinetics proposed by Nicol and Lazaro [
5] in which the maximum rate of dissolution is proportional to the initial concentration of H
+. The non-oxidative dissolution of chalcopyrite also seems to be promoted by temperature.
3.2. Rest Potentials
The initial solution was saturated with 0.5 M H
2S. Fe(III) and Cu(II) were added as Fe
2(SO
4)
3·xH
2O and CuSO
4·5H
2O respectively individually or in combination to achieve final concentrations of 10 mM and 1.75 mM, respectively. In these tests, HClO
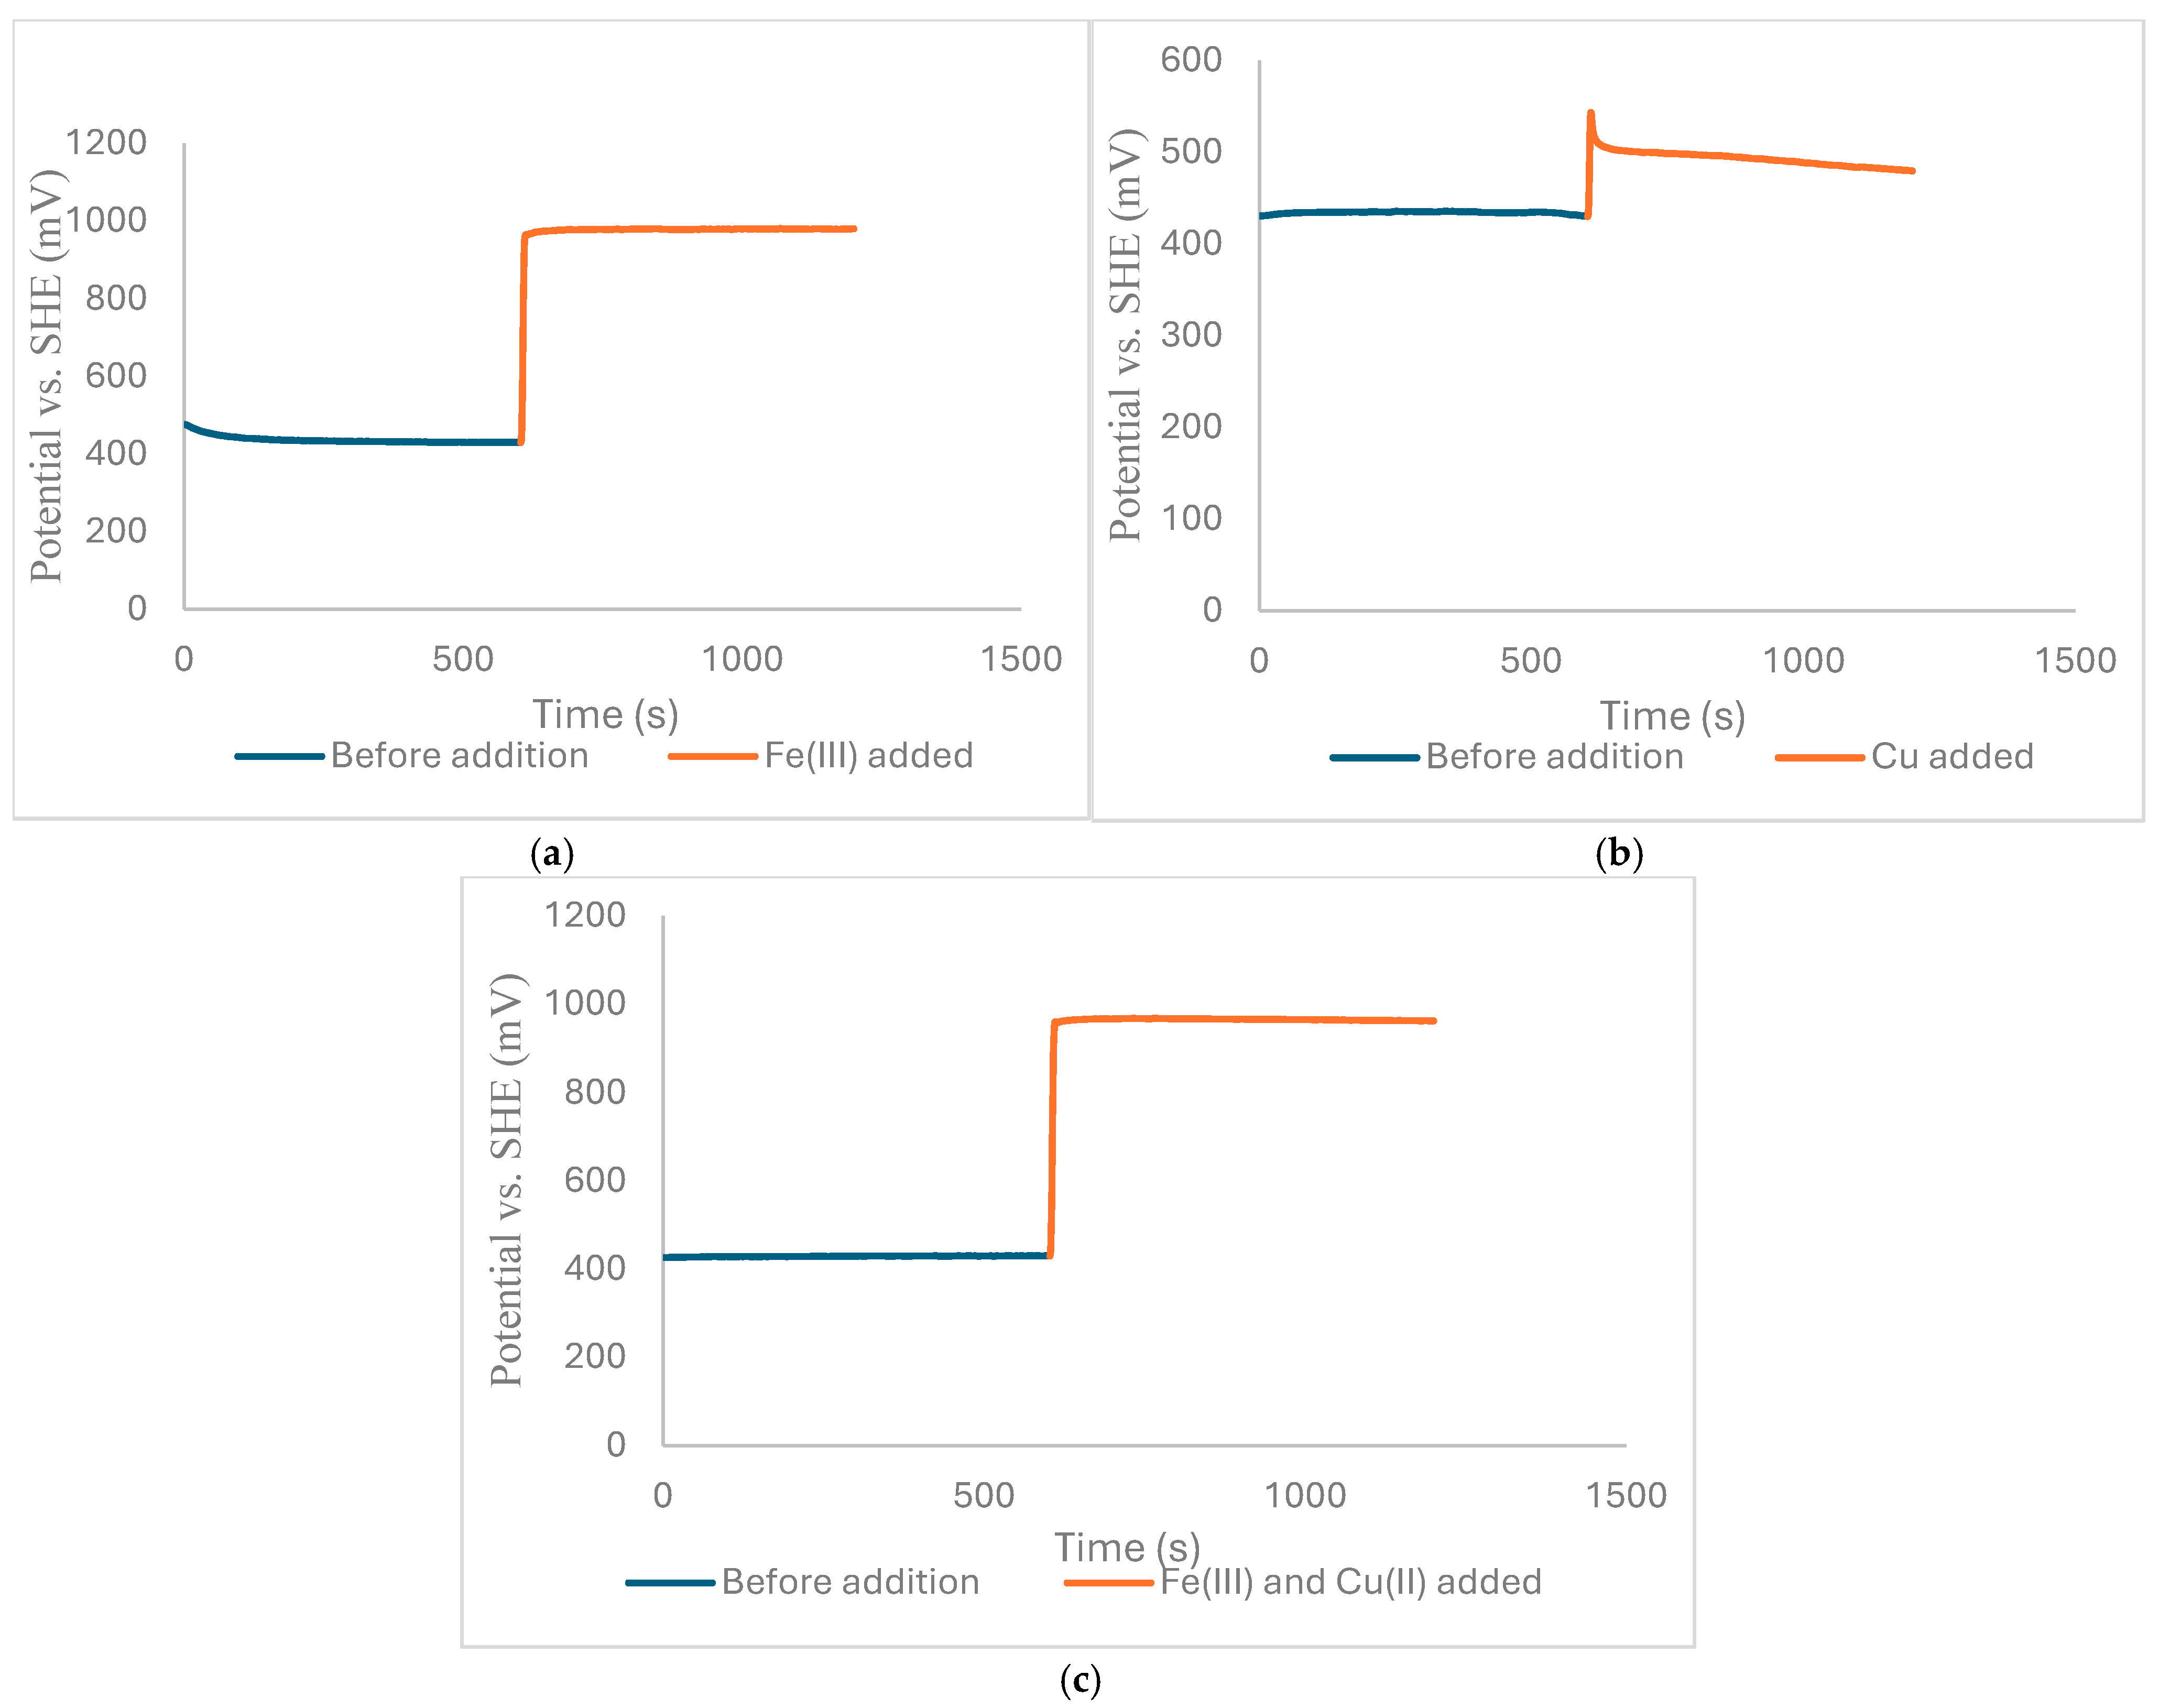
4 was added to control the pH, and it was present at the same concentration in both the initial and added solutions. At pH 2, the addition of Fe(III) (
Figure 3a), Cu(II) (
Figure 3b), or both (
Figure 3c) led to a step increase in potential. After the initial step increase, the potential remained stable in the case of Fe(III) addition (
Figure 3a) and showed a slight decrease but quickly stabilized in case of simultaneous addition of Cu(II) and Fe(III) (
Figure 3c). However, the addition of Cu(II) alone (
Figure 3b) showed a slow but noteworthy gradual decrease of the potential after the initial step increase. This decrease in potential suggests possible formation of oxidizable species, possibly the likes of Cu
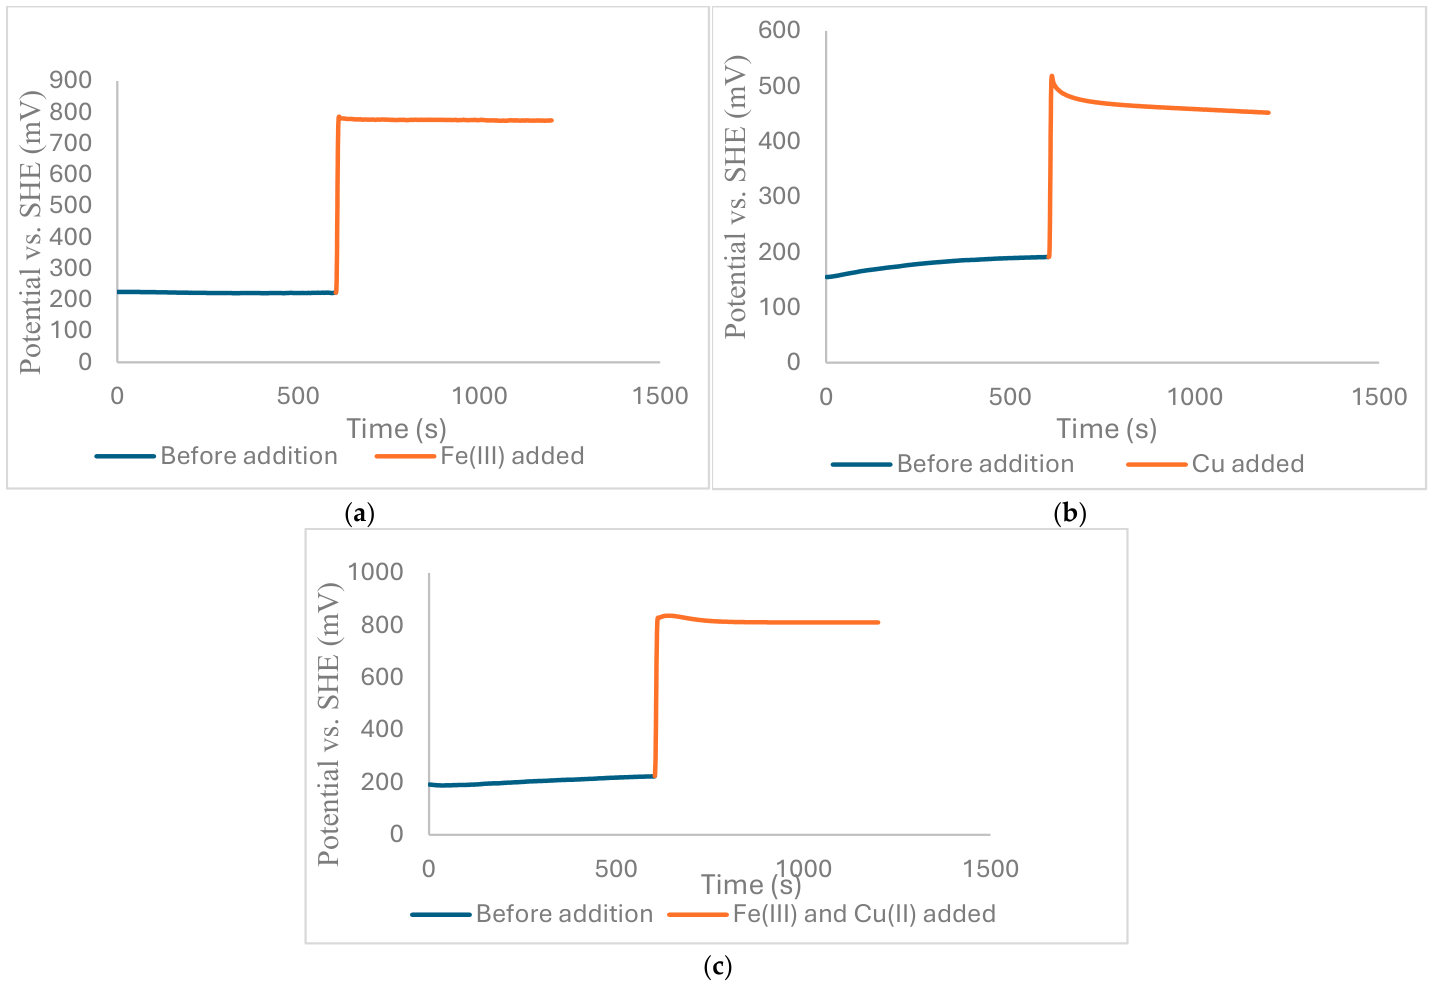
2S, at the electrode surface or in the solution. At pH 0 (
Figure 4), similar results were observed except in the case of simultaneous addition of Cu(II) and Fe(III), which led to a potential decrease after the initial step increase at pH 2 but not at pH 0.
3.3. Cyclic Voltammetry
The results observed over time when Fe(III), Cu(II), or both ions were added in acidic solution of H
2S at pH 2, with HClO
4 controlling the pH, are shown in
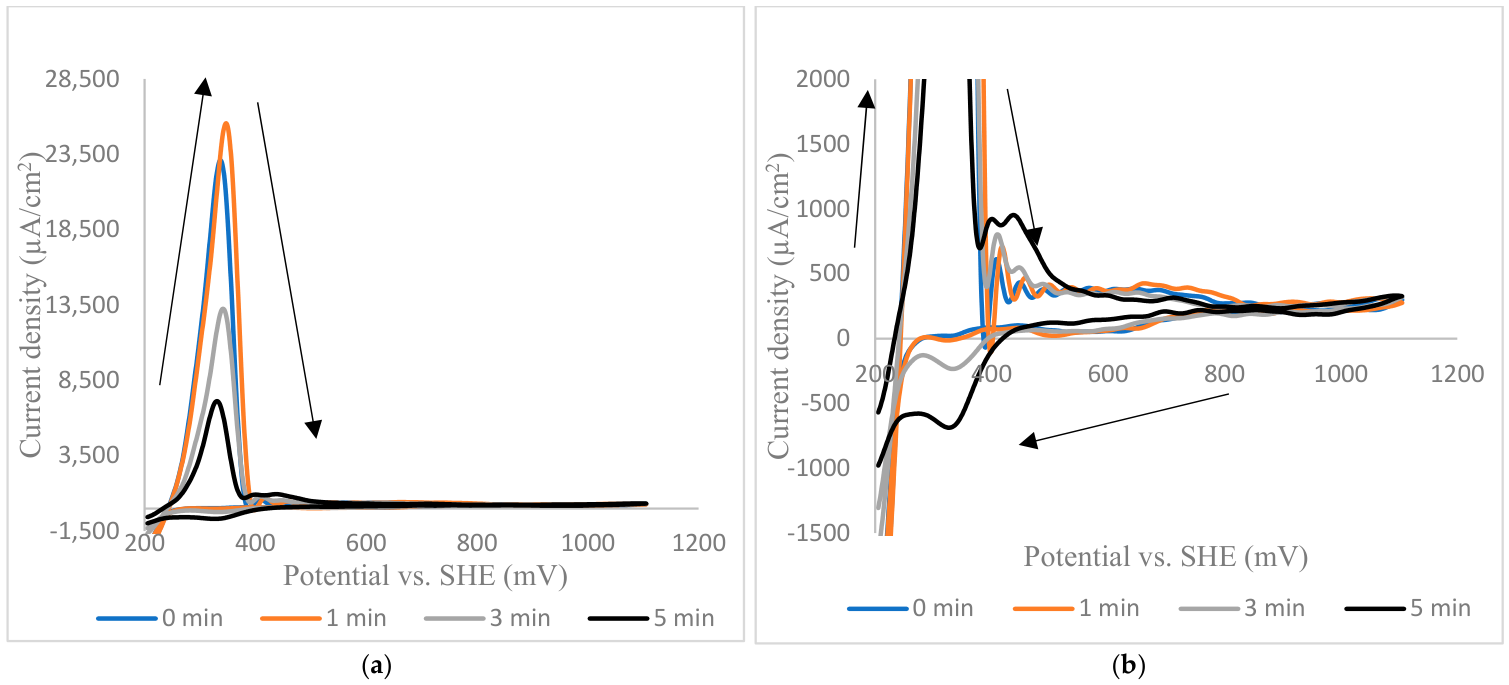
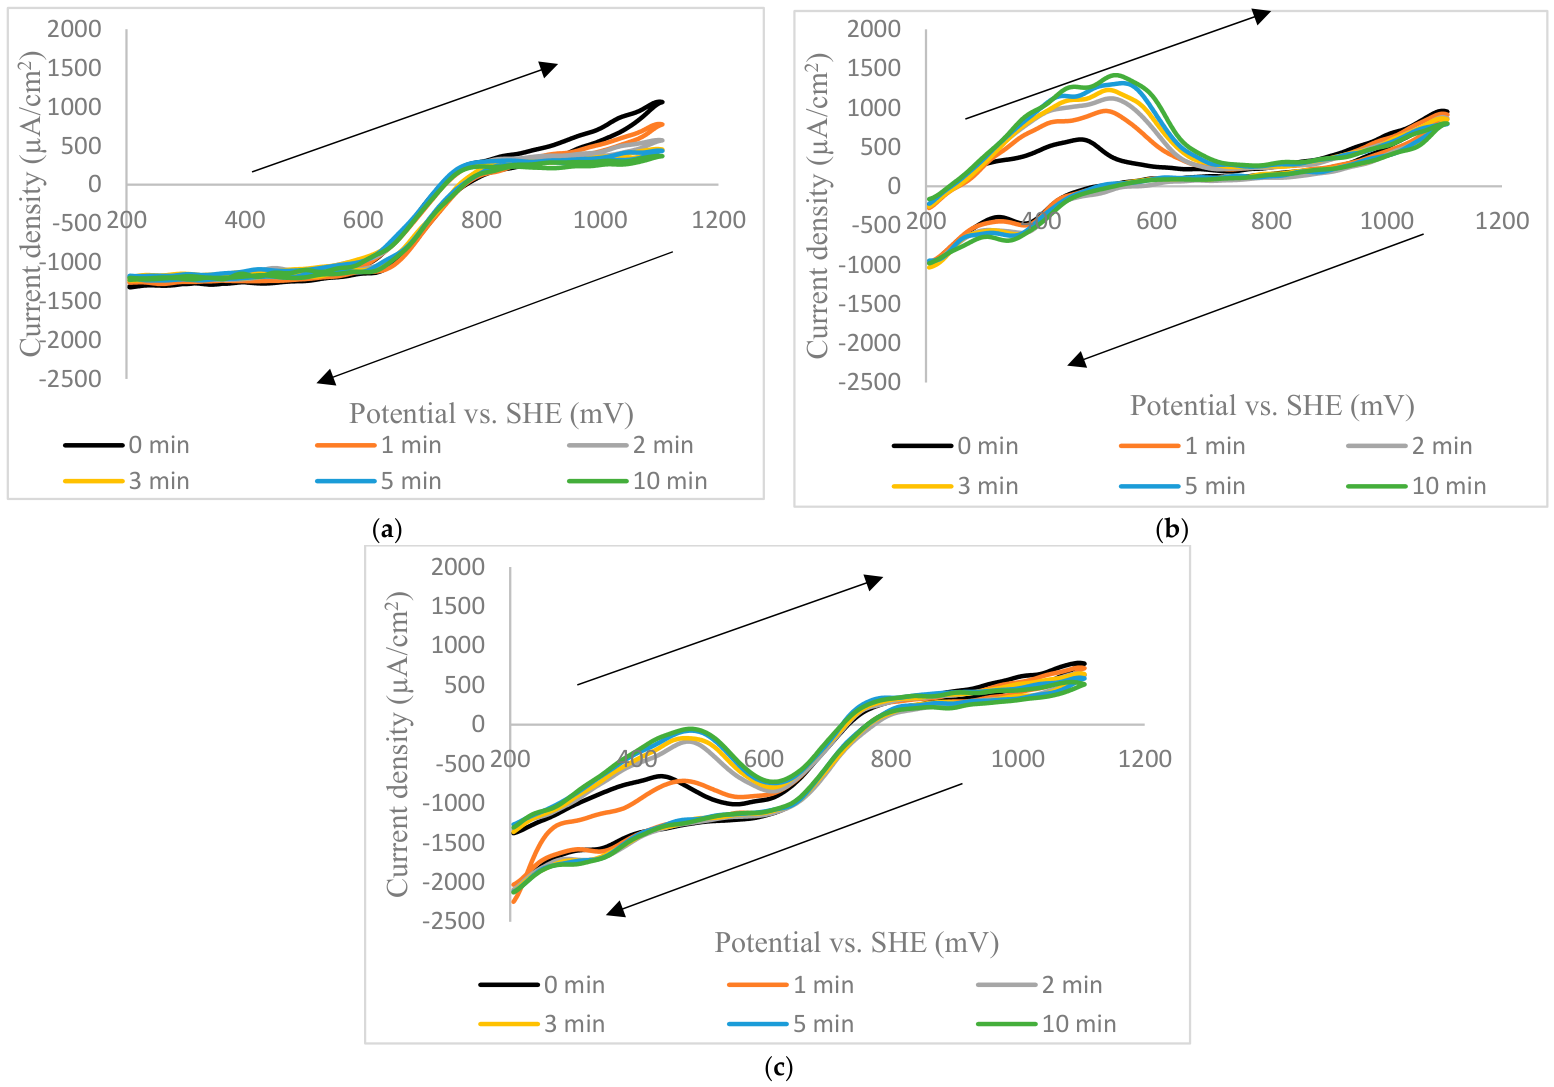
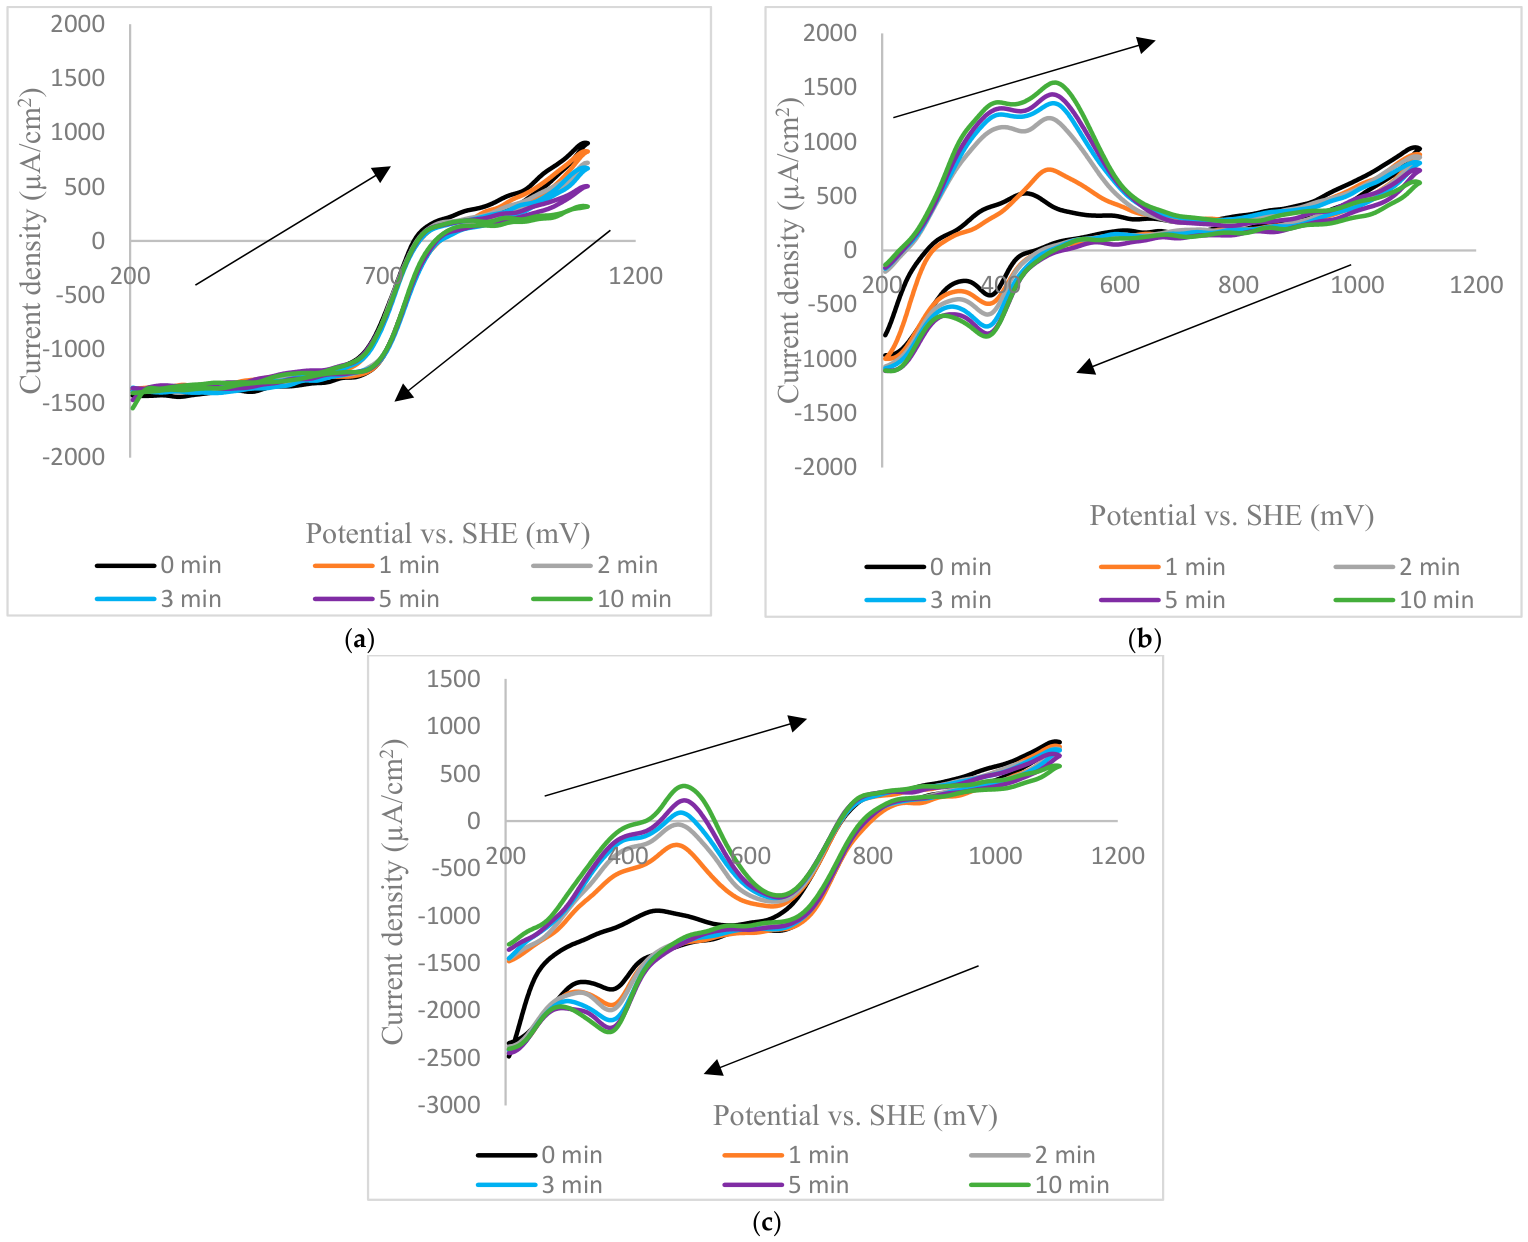
Figure 5. With the addition of Fe(III) (5 mM) (
Figure 5a), currents were negative below 800 mV during the cathodic sweep, likely due to Fe(III) reduction, and no peak was observed; rather, the curves plateaued suggesting mass transfer limited transport of Fe(III) to the mineral surface. With the addition of Cu(II) (1.75 mM) (
Figure 5b), a peak appeared around 400 to 500 mV when the electrode was scanned from 205 to 1105 mV. This peak was accompanied by a much smaller peak appearing during the reverse scan. The intensity of both peaks increased as time progressed. This peak was likely due to the oxidation of Cu(I) species, which must have formed in solution. The increase of its intensity as time progressed suggests the increase in concentration of surface compounds on the electrode. The peak observed during the reverse scan at around 300 to 400 mV is related to Cu(II) reduction, most likely to Cu(I). With the addition of both Cu(II) and Fe(III) (
Figure 5c), the current densities were below 800 mV in the cathodic sweep and the peak at 400–500 mV shifted toward increasingly less negative current density values, indicating a competing anodic reaction over and above the contribution of Fe(III) reduction currents. Even in this case, there was a local maximum that mirrored the peaks observed when Cu(II) was added alone, both in terms of peak location and trend, as the time progressed. This indicates that the Cu(I) species responsible for the peak still forms and may remain metastable despite the presence of Fe(III).
The current responses observed between 900 and 1050 mV (
Figure 5) are due to H
2S oxidation [
11,
12,
13]. Their intensity decreased with time and thus pointed to a decrease in H
2S solution concentration, due to its oxidation by Fe(III). Nevertheless, above 800 mV, the presence of Cu(II) did not result in observable impacts on the current densities. This established that Cu(II) did not catalyze H
2S oxidation by Fe(III).
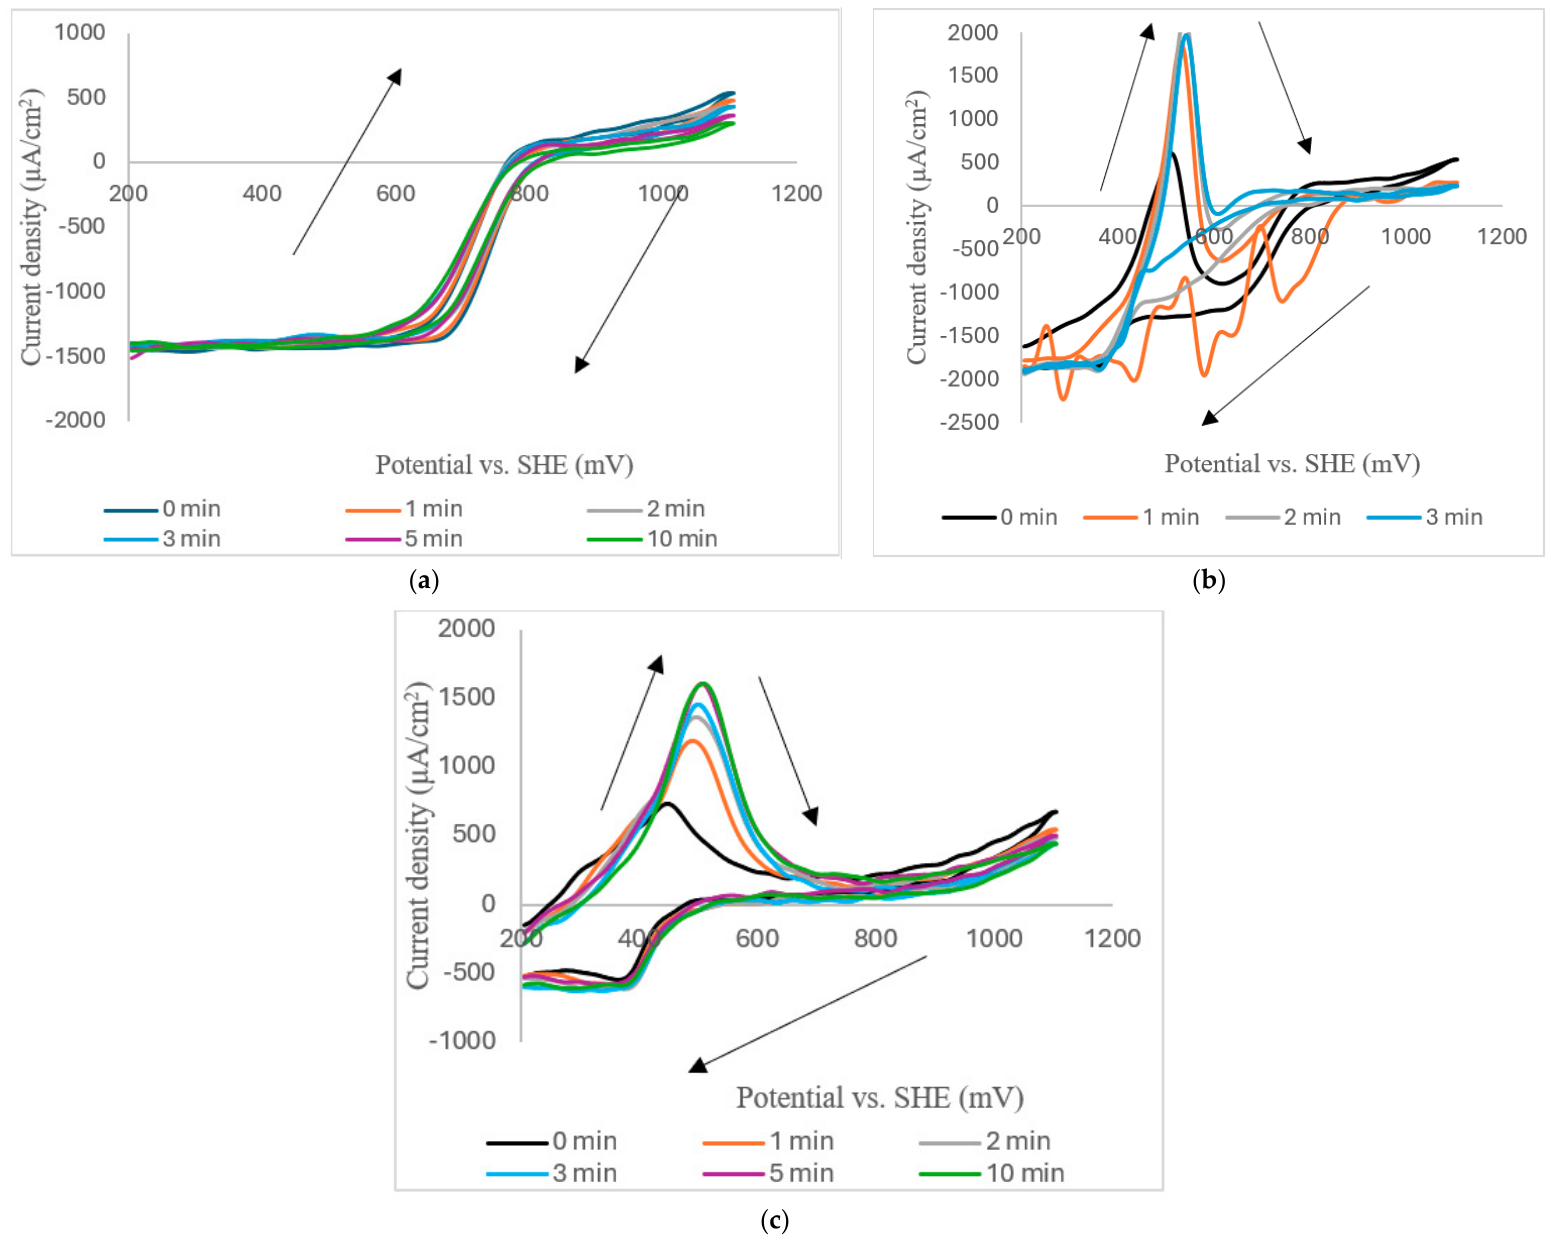
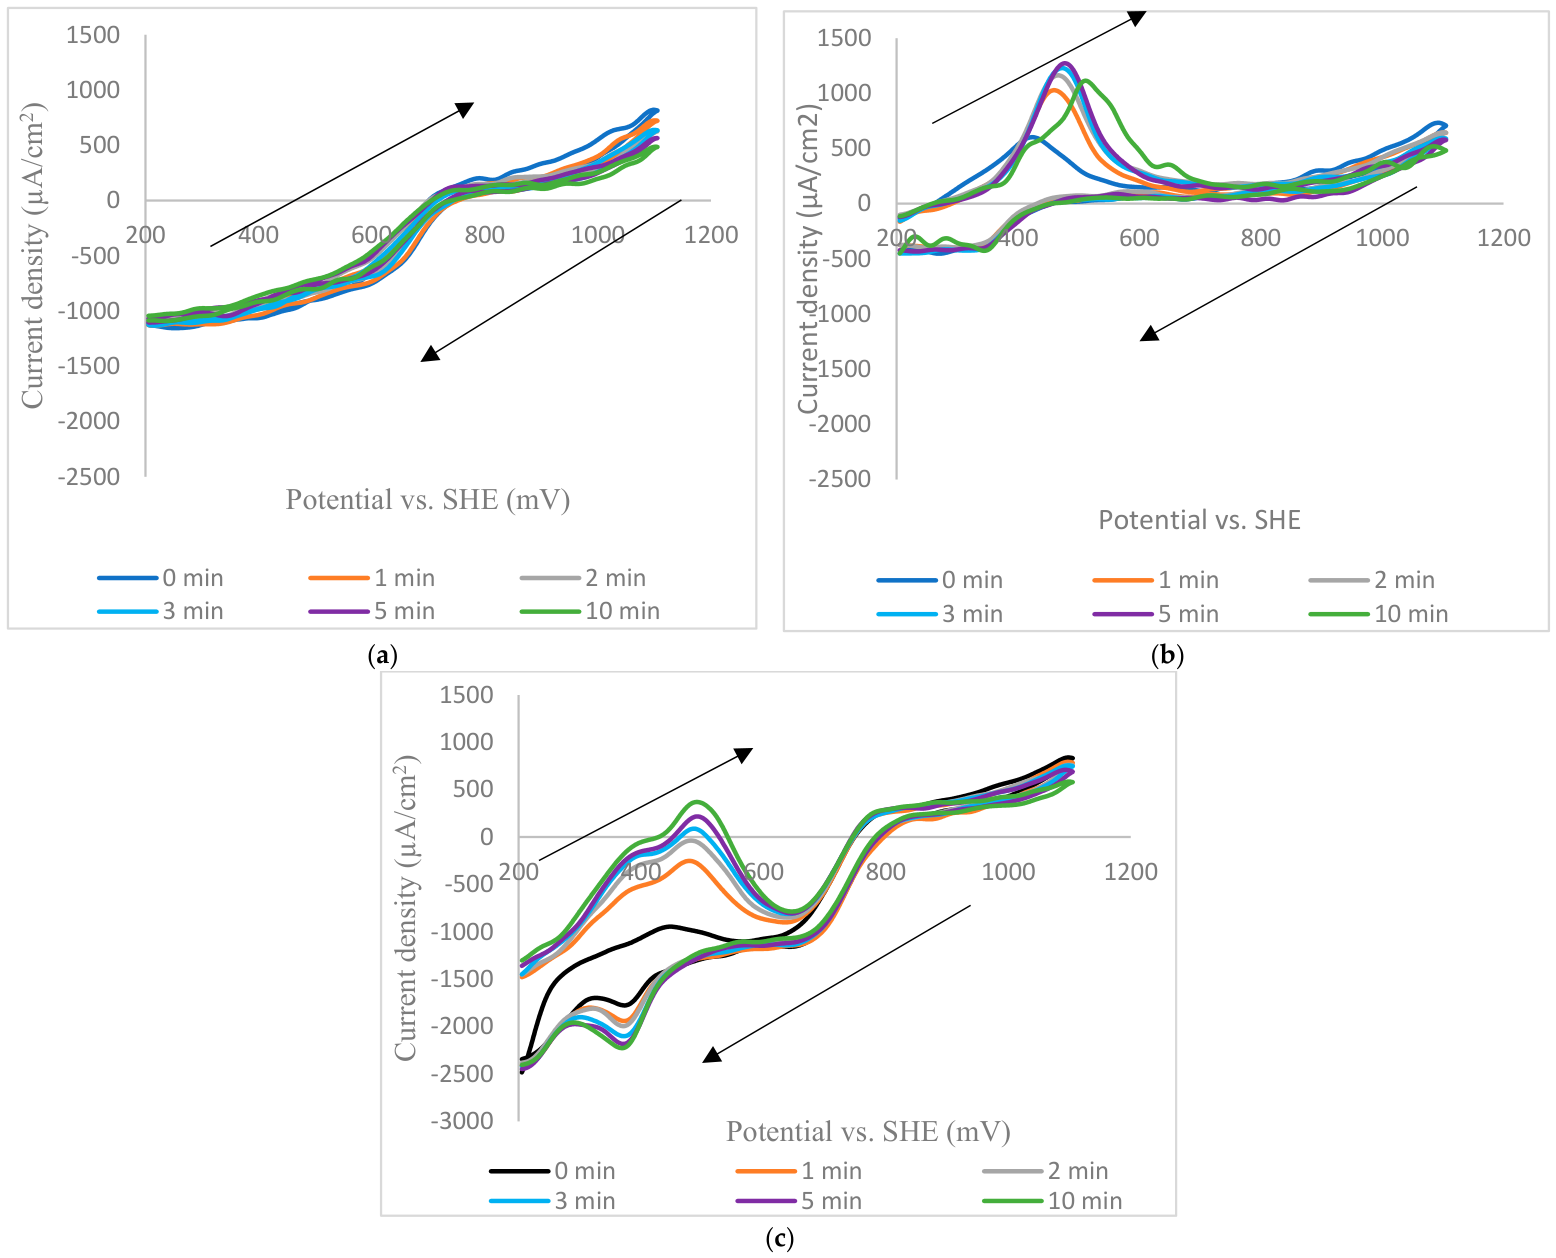
The same results were observed at pH 0 and 1 with HClO
4 and at pH 0 with H
2SO
4 (respectively
Figure 6,
Figure 7 and
Figure 8) with either 1.75 mM or 8.75 mM Cu(II) concentration. The only exception was for the addition of both Fe(III) and Cu(II) (8.75 mM) at pH 0 with HClO
4 where the peak intensity after 5 min was higher than when Cu(II) was added alone. Additionally, the peak intensity after 5 min was higher than after 10 min.
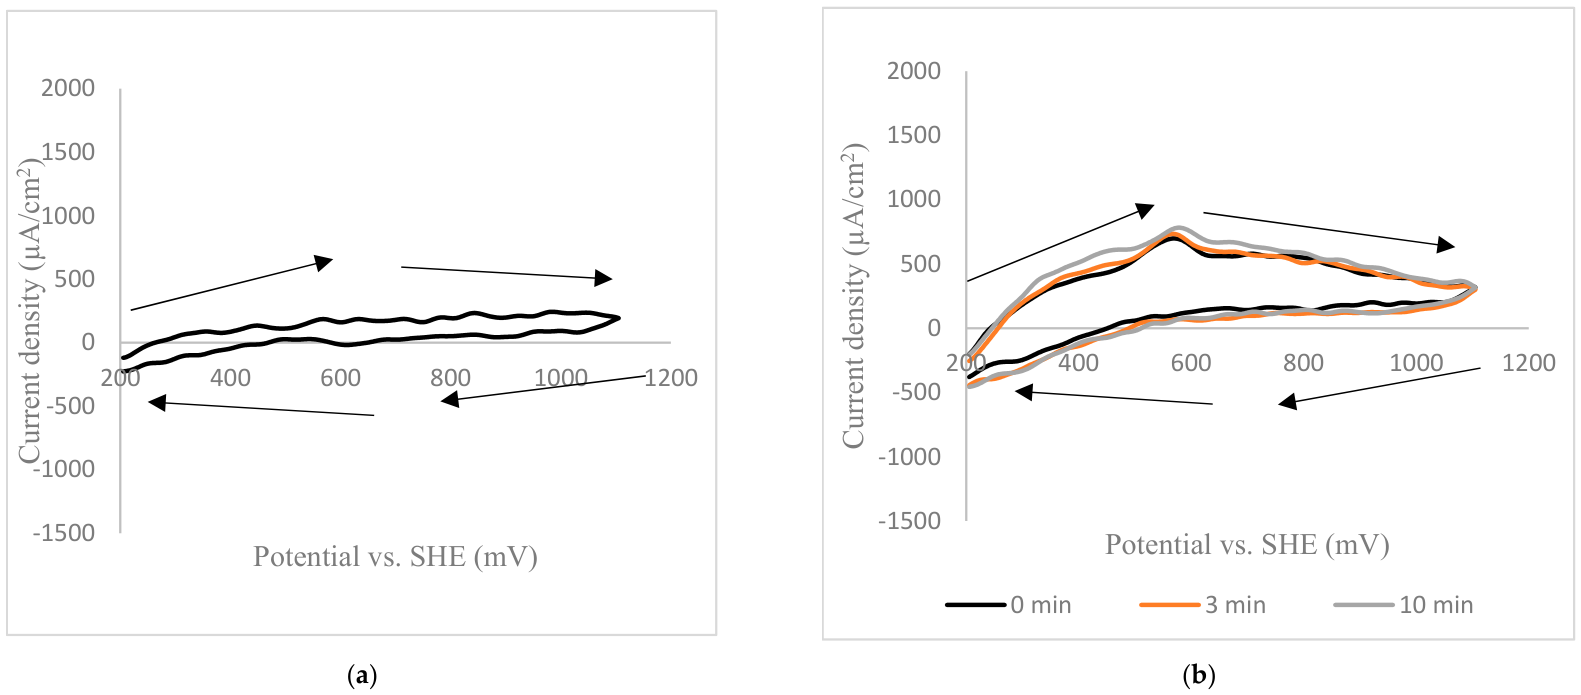
In the absence of both H
2S and the acids, no peak was observed with a solution containing 3.5 mM of initial Cu(II) (
Figure 9a). Similarly, the addition of Cu(II) at an even higher concentration (8.75 mM) to an aqueous solution without H
2S and acids led to a barely noticeable peak around 500 to 600 mV and its intensity did not significantly change as time progressed (
Figure 9b). This subtle peak may be noise or the re-oxidation of Cu(0) back to Cu(II). The area under the peak after 10 min corresponded to a charge of 188 µC. In comparison, in
Figure 7b, the corresponding charge was approximately 600 µC after 2 min and in
Figure 7c, it attained a value of 430 µC after two minutes. Importantly, in
Figure 9b, there was no corresponding cathodic peak during the reverse sweep. Therefore, it is reasonable to infer that the peak observed during the reverse sweep after the addition of Cu(II) in the presence of H
2S was related to the formation of a Cu(I) sulfide.
The addition of Cu(II) (8.75 mM) to the acid solution (1 M HClO
4, pH 0) in the absence of H
2S resulted in no observable peak in the range 400 to 600 mV (
Figure 10), suggesting the peak observed earlier was due to the presence of a Cu and S compound. However, an intense peak whose intensity decreased with time was observed between 300 and 400 mV, which was confirmed in a repeat experiment. Based on the literature [
14,
15,
16,
17,
18], this peak can be attributed to the anodic dissolution of hydrogen (Equation (13)) adsorbed on the surface of the Pt electrode through an underpotential deposition process (UPD) (Equation (14)). It was accompanied by a small peak appearing at 400 to 500 mV that became more apparent after 5 min. This peak is likely associated with the oxidation of Cu(I). The absence of the UPD-related peak when H
2S was in solution can be explained by the higher affinity of sulfur compounds for platinum. This may favor the adsorption of H
2S over H
+ on the electrode surface.
3.4. Role of Cu(II) in the Non-Oxidative/Oxidative Mechanism of Chalcopyrite in Sulfate Media
So far, results from this study have shown that H
2S formed through non-oxidative leaching of chalcopyrite can react with Cu(II) to form a Cu(I) sulfide species in sulfate and perchlorate media and in the absence of Cu(I) complexing agents. This is consistent with the mechanism proposed by Nicol et al. [
6] in chloride media (Equations (7) and (8)). While a Cu(HS)
+ complex was not directly observed during the experiments, the Cu(I) sulfide species observed could be Cu(H
2S)
+ or alike. This Cu(I) complex can be oxidized by O
2 (Equation (14)) and thus act as an intermediary in chalcopyrite leaching through the so-called non-oxidative/oxidative mechanism in sulfate media. This explanation can help better understand the positive effect of Cu(II) on chalcopyrite leaching in sulfate system media reported by previous studies [
1,
2,
3,
4].
3.5. Oxidation of H2S by the Acids
Nicol and Lazaro [
5] proposed that H
2S formed during the non-oxidative step could be later oxidized by Fe(III). Nicol et al. [
6] proposed that oxygen can oxidize H
2S at an even faster rate than Cu(II) in the chloride system. However, in the current study, evidence showed that under strongly acidic conditions, either H
2SO
4 or HClO
4 could fulfil the role of the oxidant for H
2S even at ambient temperature. The mixing of either acid (0.99 M H
2SO
4 or 1 M HClO
4 to have pH 0) with NaSH (0.025 M) resulted in the solution becoming cloudy and white (experiments whose results are reported in
Figure 6 and
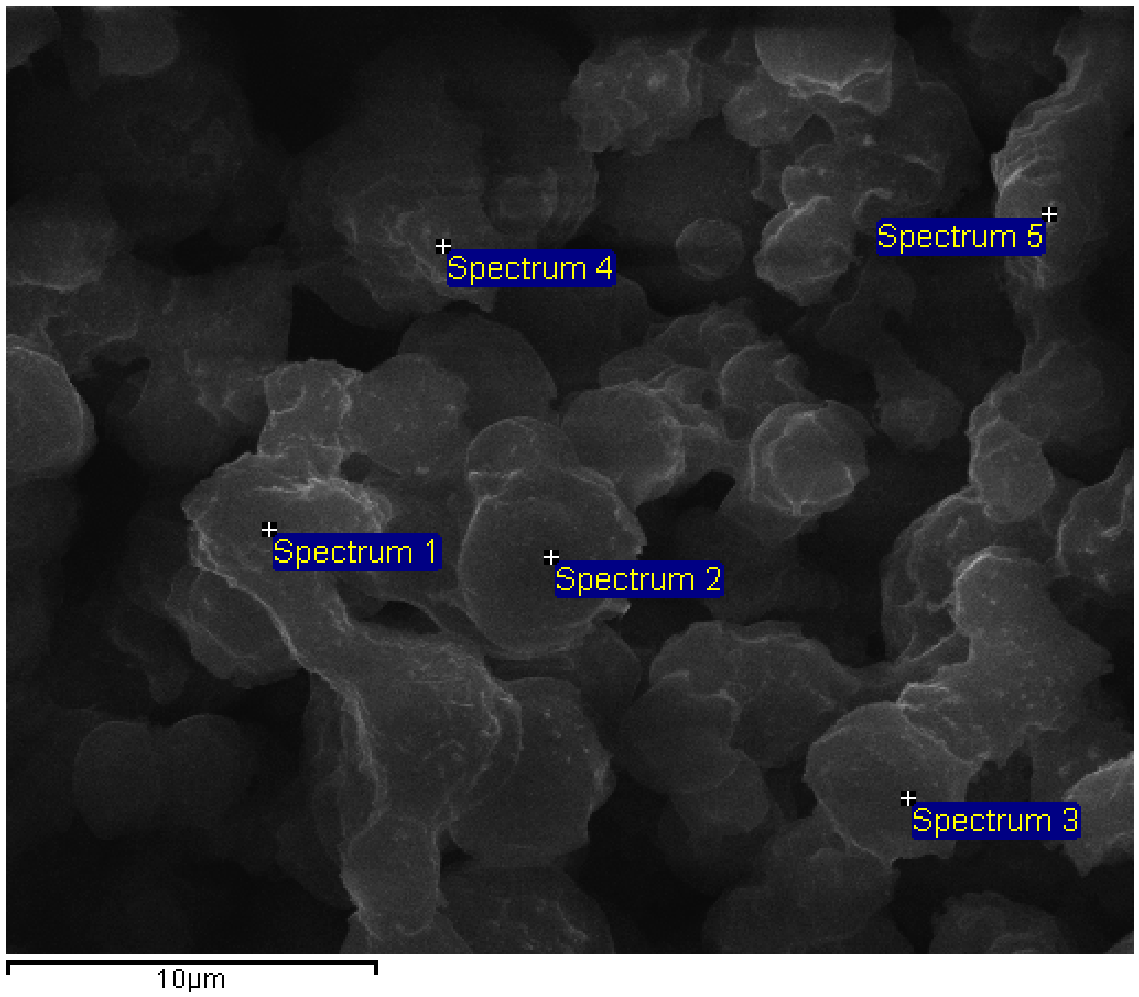
Figure 8). Over time, a yellow precipitate formed. The precipitate was further analyzed by SEM-EDS. The results (
Figure 11 and
Table 2) showed it was composed entirely of sulfur. This could be elemental sulfur formed by the oxidation of H
2S by the acids (more accurately by chlorate and sulfate anions, as per Equations (15) and (16)) or polysulfides, or even both.
Equations (17)–(19) [
19] show that both ClO
4− and SO
42− can oxidize H
2S but SO
42− is a much weaker oxidant. Zhang et al. [
20] reported that H
2S can be oxidized by H
2SO
4. The reaction was studied at 120 °C, but in their thermodynamics study the reaction was shown to be possible at even lower temperatures (50 °C) for an equimolar ratio of H
2S and H
2SO
4. Therefore, at a much larger excess of H
2SO
4 and HClO
4 as the one used in the above-mentioned experiment (about 40 times), it should not be surprising that H
2S undergoes oxidation even at room temperature.
Therefore, in sulfate and perchlorate media, oxidation of H2S formed on the surface of chalcopyrite by SO42− (Equation (15)) cannot be excluded.
3.6. Leaching Tests
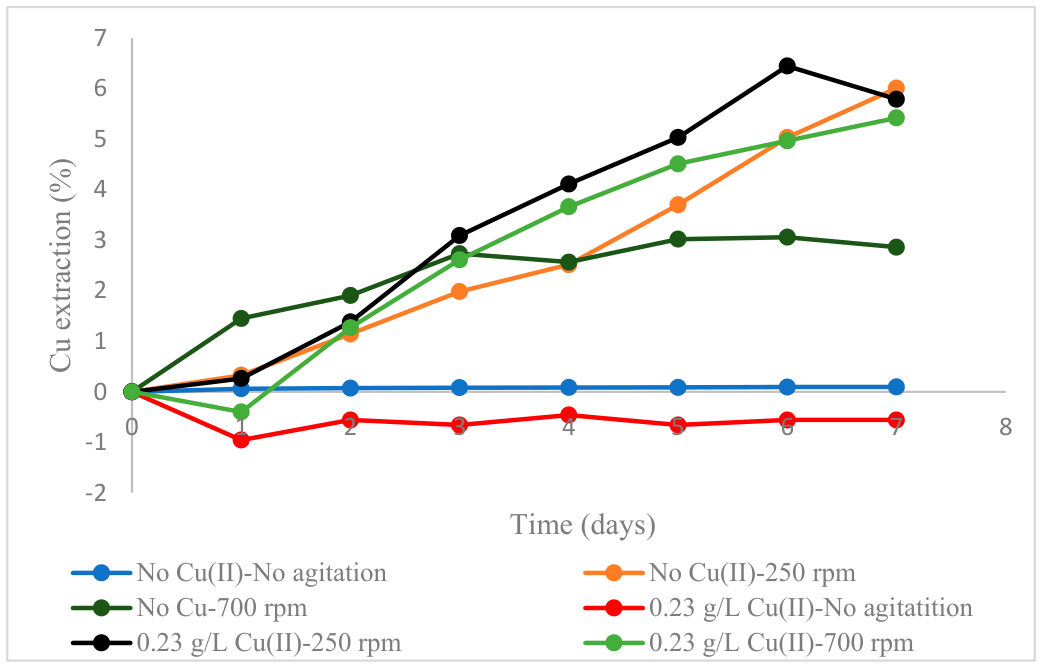
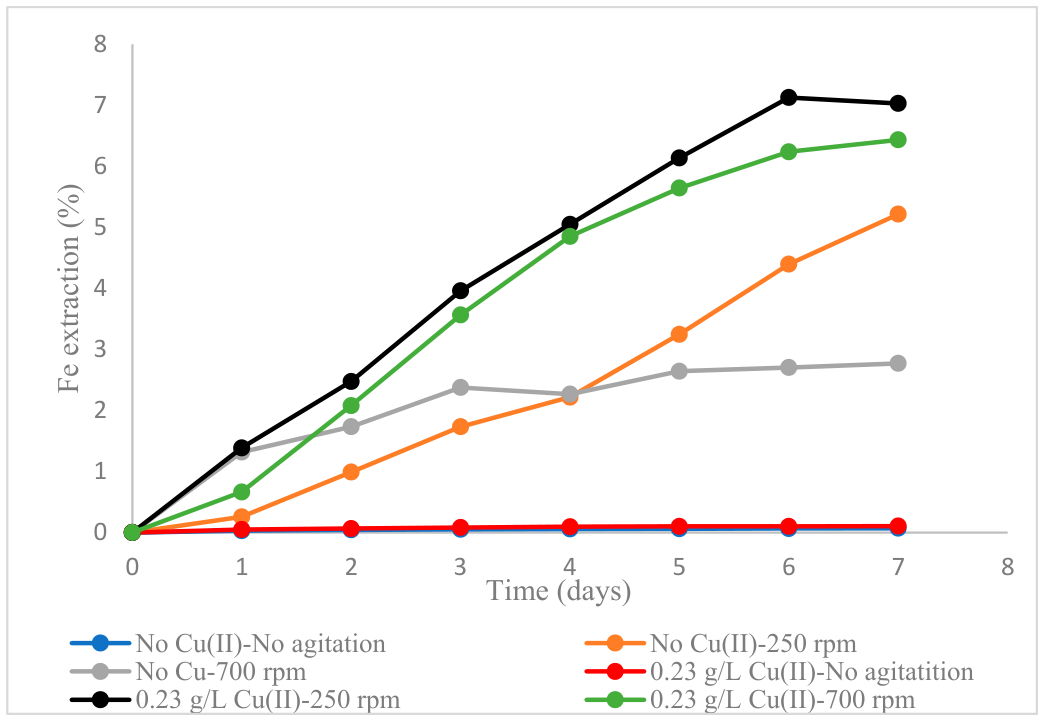
It should noted that the tests were performed as exploratory single run tests. Although the instrument error was very small (RSD <2%), the global uncertainty of 5 to 10% should be considered for the curves and the results should be seen as general kinetics trends. The Cu and Fe extractions observed under passive aeration and with 0.23 g/L (3.6 mM) initial Cu (II) at different agitation rates are presented in
Figure 12 and
Figure 13. Under nitrogen, Cu extraction was insignificant, below 1% in the absence of initial Cu [
21]. Notably, in tests where initial copper was present, concentrations declined, suggesting copper precipitation. Similarly, very little Fe was dissolved under both N
2 and passive aeration, with less than 1% extracted in each of the tests. Importantly, even though the concentrations were low, even under N
2 conditions, Fe was detected in solution, in contrast to Cu under similar conditions. Overall, some dissolution appeared to be taking place under both N
2 and passive aeration, albeit to an extremely low extent. The detection of Fe in solutions that showed a regression in Cu below what was in the initial solutions (solutions with 0.23 and 0.5 g/L initial Cu under nitrogen) further suggests that CuFeS
2 was dissolving while Cu(II) was removed from the solution. Additionally, the observation of this dissolution under N
2 points to some form of non-oxidative dissolution mechanism of CuFeS
2. Furthermore, mass transfer appears to be a factor due to the effect of agitation speed on Cu extraction.
Under passive aeration through surface aspiration, Cu extractions (
Figure 12) were highest at 250 rpm in the presence and absence of initial Cu(II). This indicates that the dissolution of chalcopyrite is positively affected by the presence of oxygen. A possible explanation would be that oxygen likely enabled the oxidative dissolution of chalcopyrite. The decline of copper concentration observed in the presence of initial Cu(II) but absence of agitation points towards the formation of Cu precipitates regardless of the presence of oxygen.
The lack of agitation likely limited oxygen migration to the mineral surface, which, combined with its poor solubility in water, resulted in the system behaving similarly to conditions under nitrogen. Agitation improved the mass transfer of oxygen or other solution species to the mineral surface resulting in higher Cu and Fe extractions in the presence of oxygen, while under nitrogen this results only in higher Fe extractions but increased Cu precipitation. Therefore, excessive agitation rate does not necessarily translate into better Cu extractions but most likely generates the release of new species by abrasion.
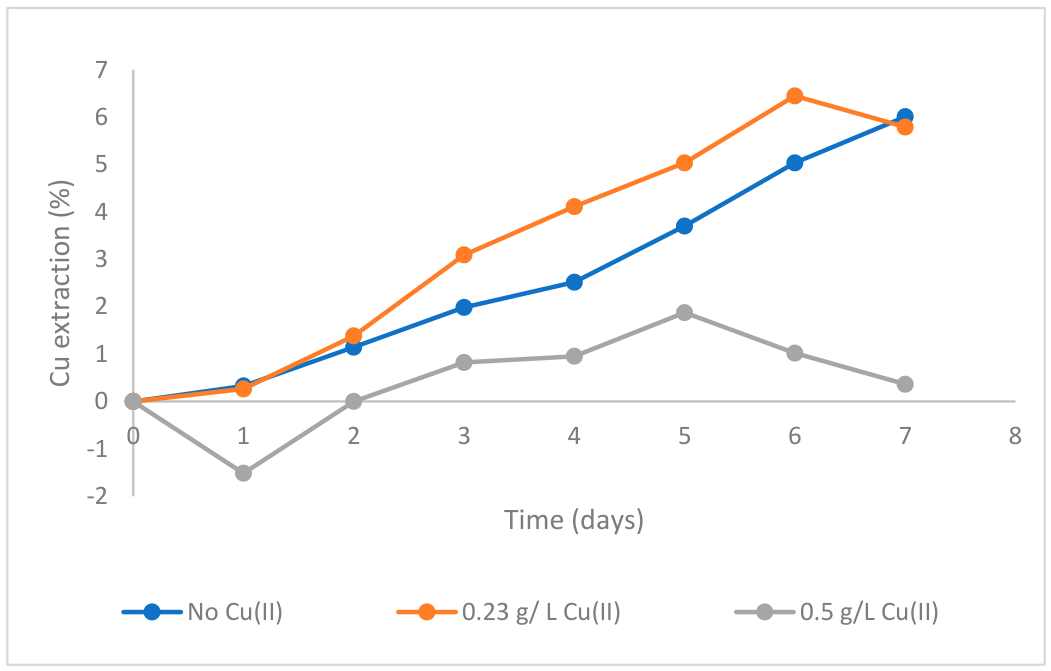
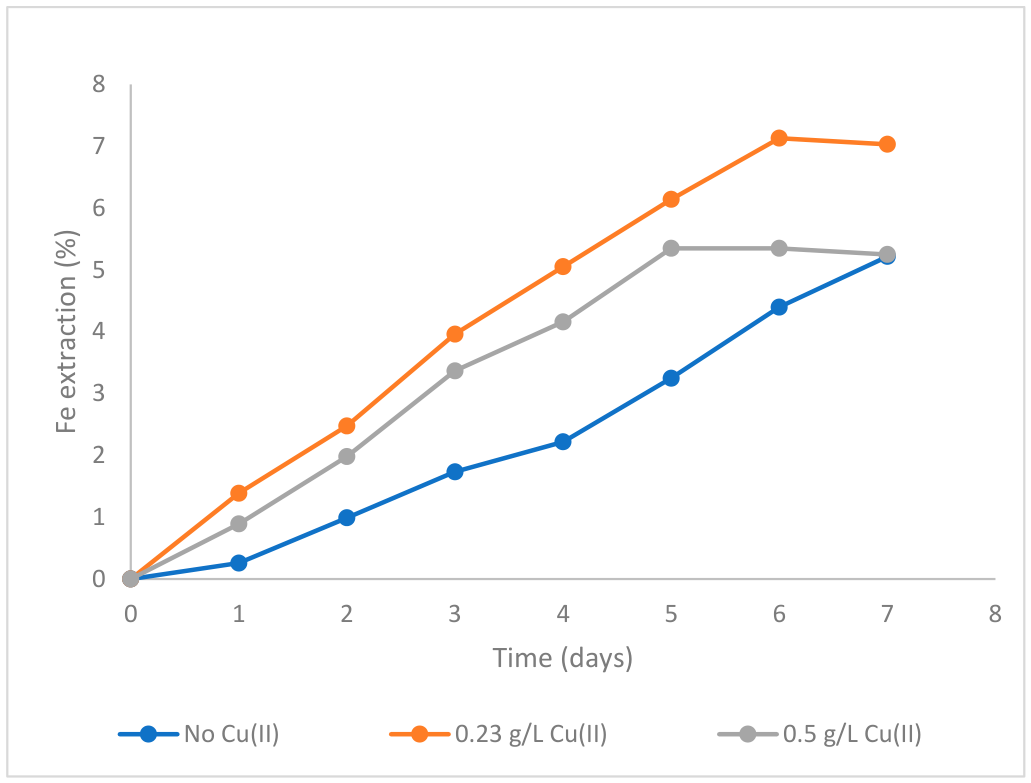
The effect on initial Cu(II) concentration was studied in the presence of oxygen at an agitation rate of 250 rpm at 0 g/L Cu(II), at 0.23 g/L Cu(II), and 0.5 g/L Cu(II). The results are shown in
Figure 14 for Cu extractions and
Figure 15 for Fe extractions.
The lowest Cu extractions were measured with 0.5 g/L initial Cu(II). A decline in Cu copper concentration in solution, representing about 10% of the initial Cu input, was observed on the first day, resulting in the “negative” extraction. There was no such decline observed at 0.23 g/L initial Cu. Higher extraction was shown for the solution with 0.23 g/L initial Cu from the third day onwards, but by the seventh day, the curve overlapped with that of the Cu(II) free solution.
The highest Fe extractions were observed in the tests that had 0.23 g/L initial Cu(II) compared to the systems with 0.5 g/L initial Cu(II) and no initial Cu(II). The extraction appeared to taper off on day 6 in the 0.23 g/L initial Cu(II) and on day 5 in the 0.5 g/L initial Cu(II). This may suggest that the chalcopyrite dissolution reaction had reached its equilibrium, or the mineral surface may have become inhibited through surface precipitation.
In the absence of initial Cu(II), Cu and Fe extraction trends were fairly similar but with Fe extractions exceeding Cu. This suggests that the presence of initial Cu(II) in leach solutions had a positive impact on chalcopyrite dissolution. However, there seemed to exist an optimum initial Cu(II) concentration, as the highest initial Cu(II) concentration did not lead to enhanced dissolution of the mineral. Similar results were reported by Nicol et al. [
6] in chloride media; albeit in the chloride system the Cu(II)/Cu(I) redox couple is thermodynamically stable. Overall, Cu extraction remained less than 10% in 7 days. This is in agreement with the well-documented kinetic behavior of chalcopyrite in sulfate media at ambient temperatures [
22,
23,
24].
3.7. Insight into the Mechanism
Evidence from both the linear and cyclic voltammetry suggests the formation of H
2S by the non-oxidative acid attack on chalcopyrite observed under nitrogen conditions, (Equation (5)), as established by Nicol and Lazaro [
5]. H
2S formed by the acid non-oxidative leaching of chalcopyrite can be oxidized by Fe(III) in sulfate media to form elemental sulfur (Equation (4)) [
5]. Nicol et al. [
6] show that O
2 does not directly oxidize H
2S in chloride media. Rather, H
2S is oxidized by Cu(II), forming Cu(I), which is later oxidized by O
2. Another possible route for the oxidation of H
2S in a highly acidic sulfuric acid solution would be through the reaction with SO
42− (Equation (16)).
The leaching results show that Cu(II) is likely to have a catalytic effect on the dissolution of chalcopyrite in the presence of O
2. At the same time, cyclic voltammetry results clearly show that Cu(II) does not catalyze the oxidation of H
2S by Fe(III). This calls for an explanation involving a more direct role of O
2. Such an explanation was proposed by Nicol et al. [
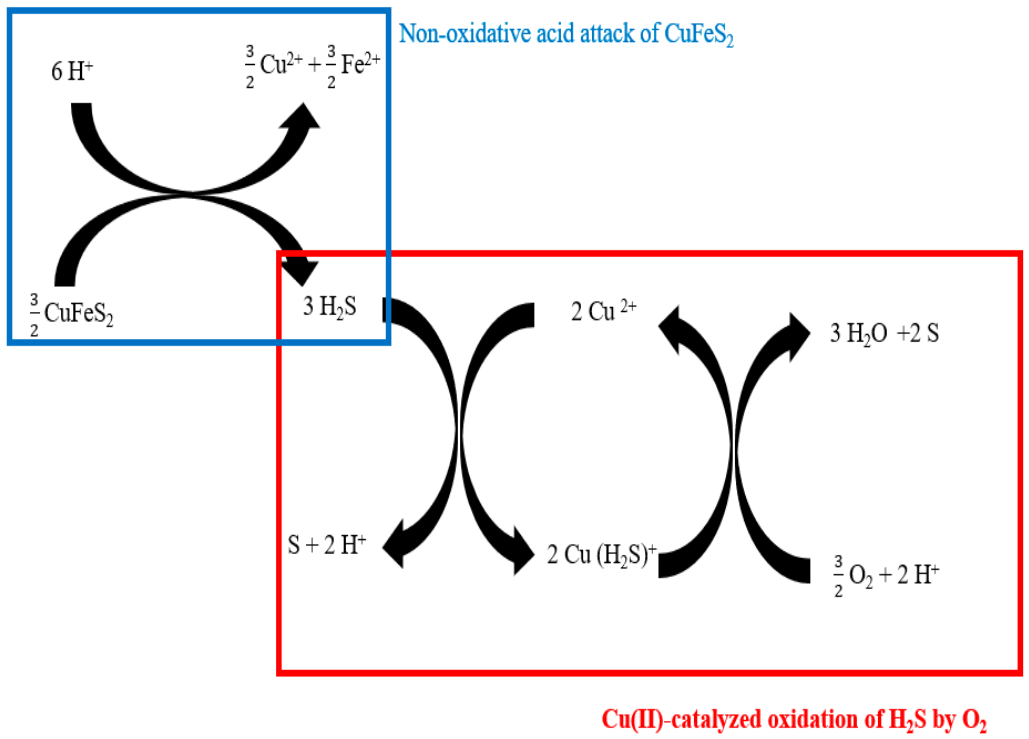
6] for the chloride media. Following the proposed mechanism, Cu(II) forms a complex with HS
− (Equation (9)), which would react faster with O
2 (Equation (10)) than uncomplexed Cu(II).
This complex Cu(HS)
+ could not be inferred from the experimental data. However, a Cu(I) species that could be Cu(H
2S)
+ or alike was observed. This complex may be the Cu intermediary, explaining its catalytic activity rather than by Cu(II), at least in sulfate media. As in the mechanism proposed by Nicol et al. [
6], the oxidation of Cu(HS)
+ by O
2 (Equation (14)) would be faster than its oxidation by Fe(III) (Equation (20)). Its oxidation by Fe(III) would be of similar or slower rate in comparison with the oxidation of H
2S by Fe(III) (Equation (4)). These hypotheses would explain both the positive effect of Cu(II) observed in the leaching tests under free-surface aspiration and in the absence of Cu(II) catalytic effect on H
2S oxidation by Fe(III) shown in cyclic voltammetry tests.
Cu(II)-catalysis of sulfur species oxidation has been reported in other studies. These include the catalysis of thiosulfate oxidation by oxygen in ammoniacal solutions [
25], in gold leaching [
26] and in acid solutions of sulfuric acid [
27]. Cu(H
2S)
+ may be the precursor to Cu precipitates like Cu
2S and CuHS via reactions (21) and (22) and even CuS. Many authors have indeed argued and shown that even in CuS, either as an amorphous precipitate [
28] or as covellite [
29,
30], Cu is found in the +1 oxidation state. The results of the leaching study suggest the formation of Cu precipitates, particularly under nitrogen. The precipitation of Cu from the reaction between soluble Cu(II) and H
2S produced by acid non-oxidative leaching has been reported by Maley et al. [
31]. Although in their study, H
2S resulted from non-oxidative leaching of pyrrhotite, H
2S produced by non-oxidative leaching of chalcopyrite can act the same way. Cu precipitation as covellite also occurs in chloride media in the absence of oxygen [
32]. This not only supports the likelihood of formation of Cu precipitates but also agrees with the leaching results.
The non-oxidative/oxidative mechanism (non-oxidative dissolution of chalcopyrite followed by oxidation of H
2S) (
Figure 16) could then also exist in a sulfate system in parallel with the “direct” oxidative dissolution of chalcopyrite. The non-oxidative/oxidative mechanism may play an important role in the early stages of the leaching process of chalcopyrite, particularly in a solution devoid of initial Fe(III) or even poor in Fe(III). The existence of an optimum Cu(II) concentration is likely related to Cu precipitation.