
Article

Carbonation Reaction Mechanisms of Portlandite Predicted from Enhanced Ab Initio Molecular Dynamics Simulations
by
Sylvia M. Mutisya
1 and
Andrey G. Kalinichev
1,2,*
1
Laboratoire SUBATECH (UMR 6457–Institut Mines-Télécom Atlantique, Université de Nantes, CNRS/IN2P3), 44307 Nantes, France
2
International Laboratory for Supercomputer Atomistic Modelling and Multiscale Analysis, National Research University Higher School of Economics, 123458 Moscow, Russia
*
Author to whom correspondence should be addressed.
Minerals 2021, 11(5), 509; https://doi.org/10.3390/min11050509
Submission received: 21 March 2021 / Revised: 4 May 2021 / Accepted: 6 May 2021 / Published: 11 May 2021
(This article belongs to the Special Issue First Principles Calculations of Minerals and Related Materials)

Abstract

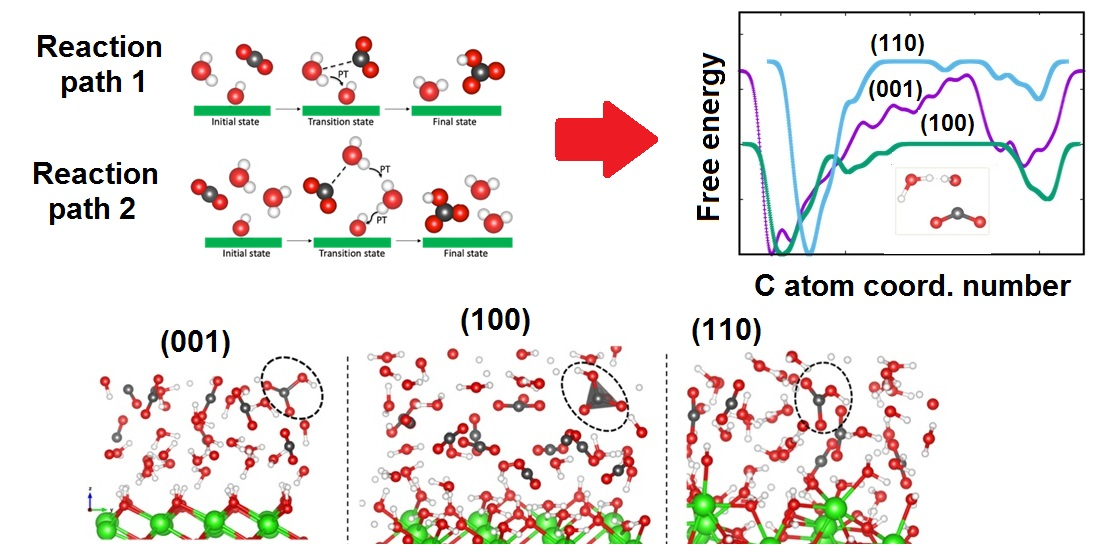
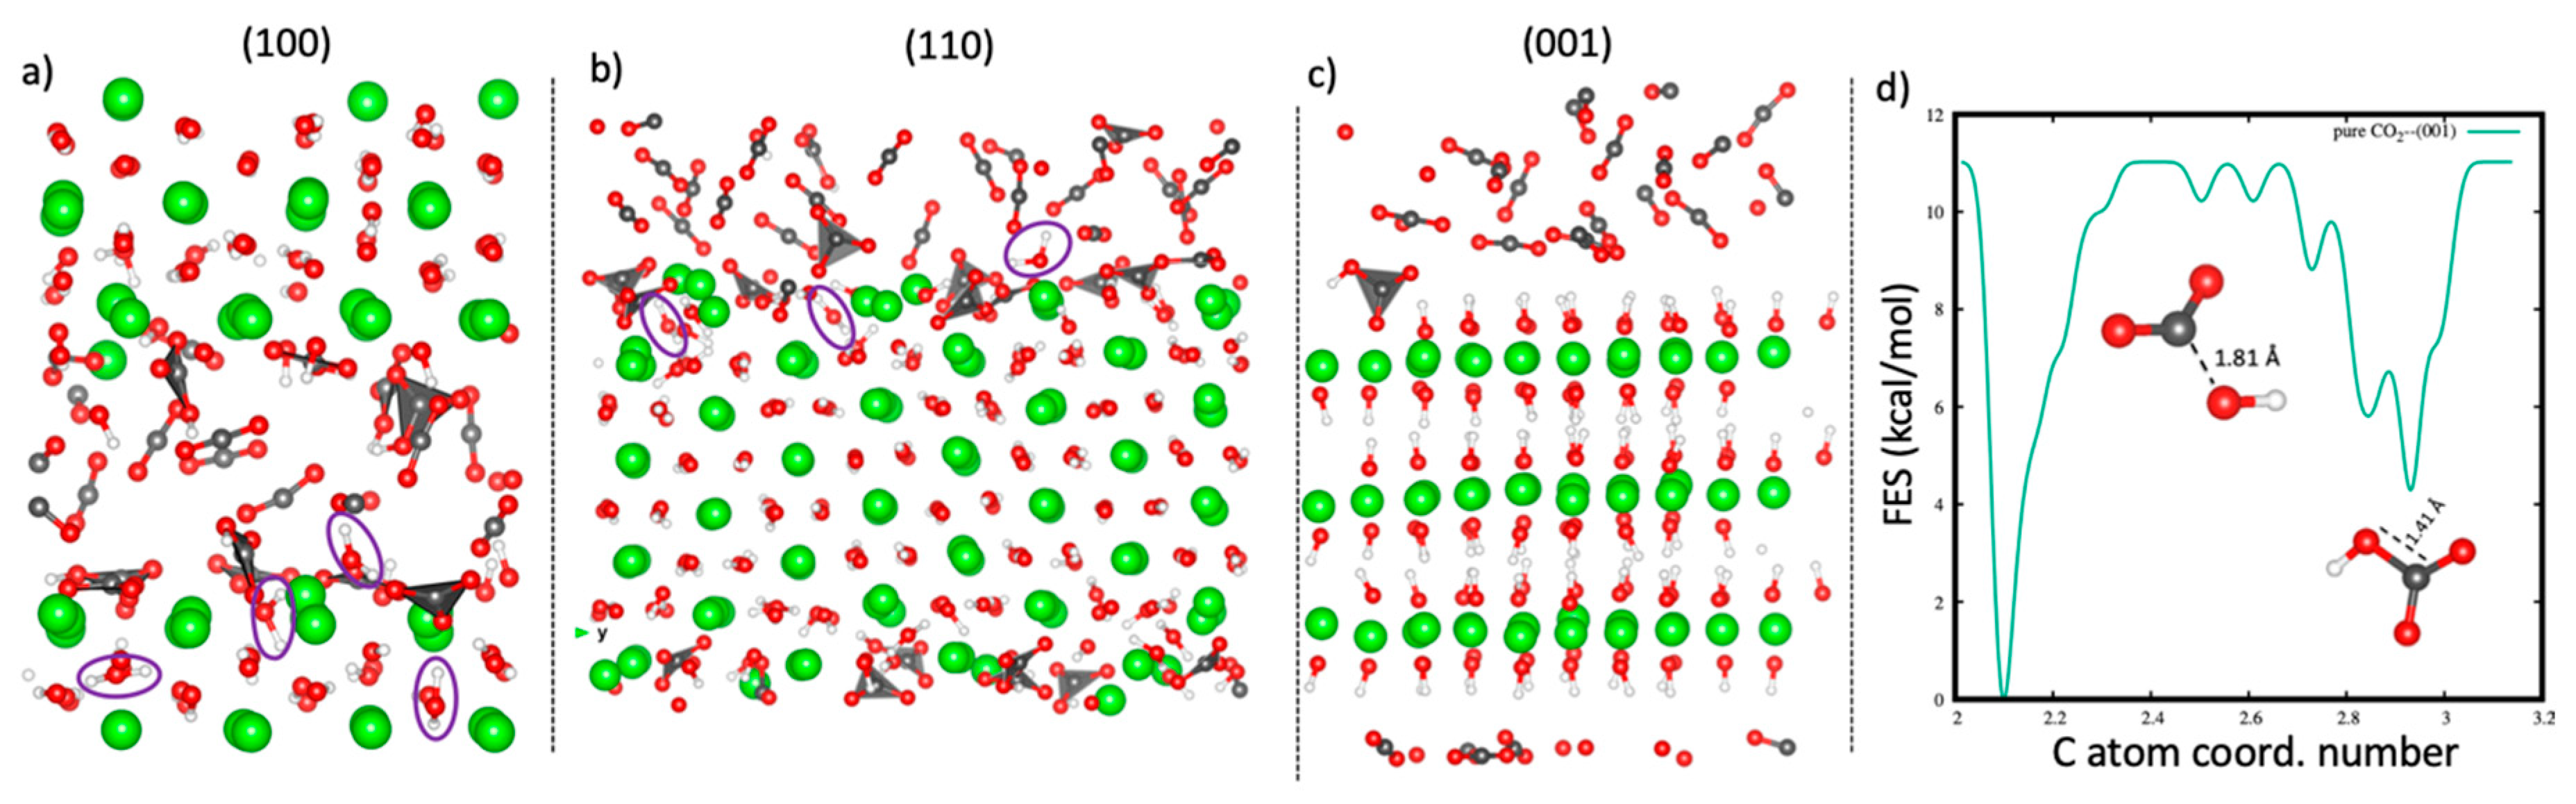
Geological carbon capture and sequestration (CCS) is a promising technology for curbing the global warming crisis by reduction of the overall carbon footprint. Degradation of cement wellbore casings due to carbonation reactions in the underground CO2 storage environment is one of the central issues in assessing the long-term success of the CCS operations. However, the complexity of hydrated cement coupled with extreme subsurface environmental conditions makes it difficult to understand the carbonation reaction mechanisms leading to the loss of well integrity. In this work, we use biased ab initio molecular dynamics (AIMD) simulations to explore the reactivity of supercritical CO2 with the basal and edge surfaces of a model hydrated cement phase—portlandite—in dry scCO2 and water-rich conditions. Our simulations show that in dry scCO2 conditions, the undercoordinated edge surfaces of portlandite experience a fast barrierless reaction with CO2, while the fully hydroxylated basal surfaces suppress the formation of carbonate ions, resulting in a higher reactivity barrier. We deduce that the rate-limiting step in scCO2 conditions is the formation of the surface carbonate barrier which controls the diffusion of CO2 through the layer. The presence of water hinders direct interaction of CO2 with portlandite as H2O molecules form well-structured surface layers. In the water-rich environment, CO2 undergoes a concerted reaction with H2O and surface hydroxyl groups to form bicarbonate complexes. We relate the variation of the free-energy barriers in the formation of the bicarbonate complexes to the structure of the water layer at the interface which is, in turn, dictated by the surface chemistry and the degree of nanoconfinement.
Keywords: cement; portlandite; carbonation; atomistic computer simulations; ab initio molecular dynamics (AIMD); metadynamics; density functional theory (DFT) cement; portlandite; carbonation; atomistic computer simulations; ab initio molecular dynamics (AIMD); metadynamics; density functional theory (DFT)
Graphical Abstract