5.2.1. Phase Behavior of the α-Olefin + Hexane + POE96k-10 System
This work examines the phase characteristics of the α-olefin +
n-hexane + POE96k-10 ternary systems, with α-olefins including C
2H
4, 1-C
4H
8, 1-C
6H
12 and 1-C
8H
16 as the third component. Among these α-olefins, C
2H
4 is the primary polymerization monomer, while 1-C
4H
8, 1-C
6H
12 and 1-C
8H
16 are widely utilized in industrial POE synthesis as comonomers. The phase transition data for the α-olefin +
n-hexane + POE96k-10 ternary systems are presented in
Table 4,
Table 5,
Table 6 and
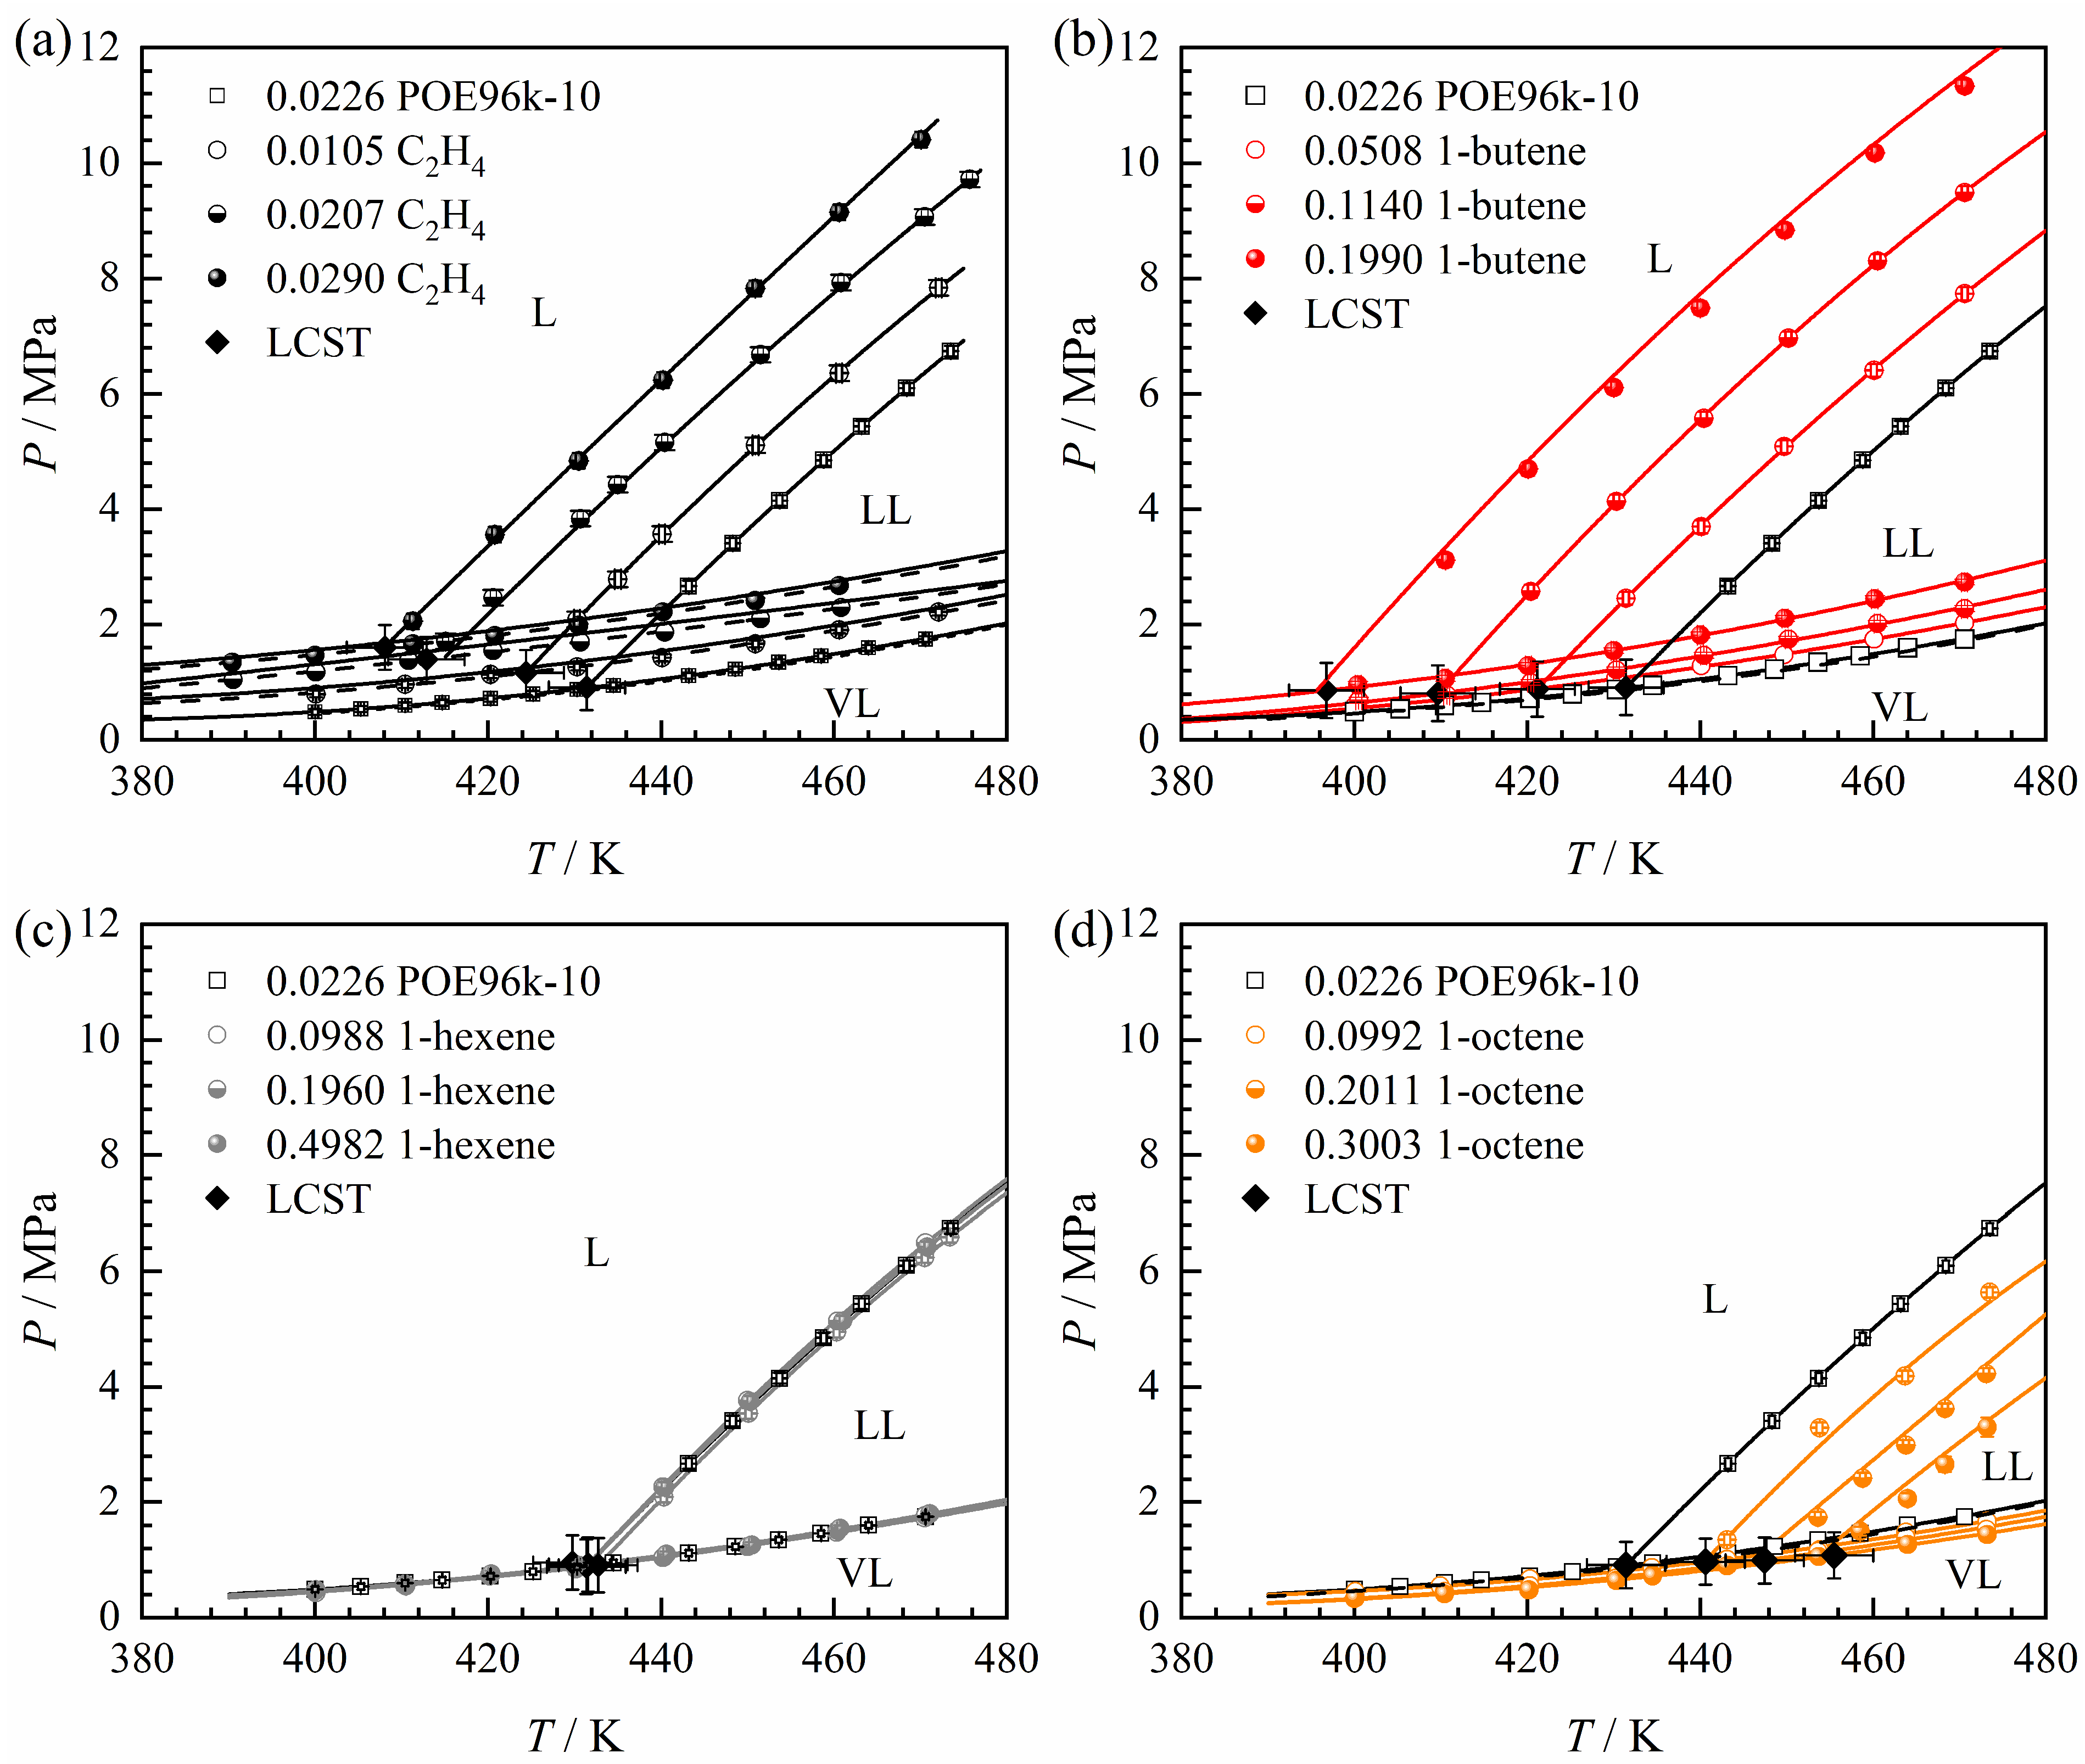
Table 7, and the phase boundaries are depicted in
Figure 5. The α-olefin content was selected according to industrial production data and is expressed relative to the solvent composition, with the polymer component excluded. To examine the distinct influence of different α-olefins on phase transitions, the
n-hexane + POE96k-10 system with a polymer mass fraction of 0.0226 g·g
–1 is used as a reference in this section while keeping the polymer mass fraction of the four ternary systems at approximately 0.023 g·g
–1.
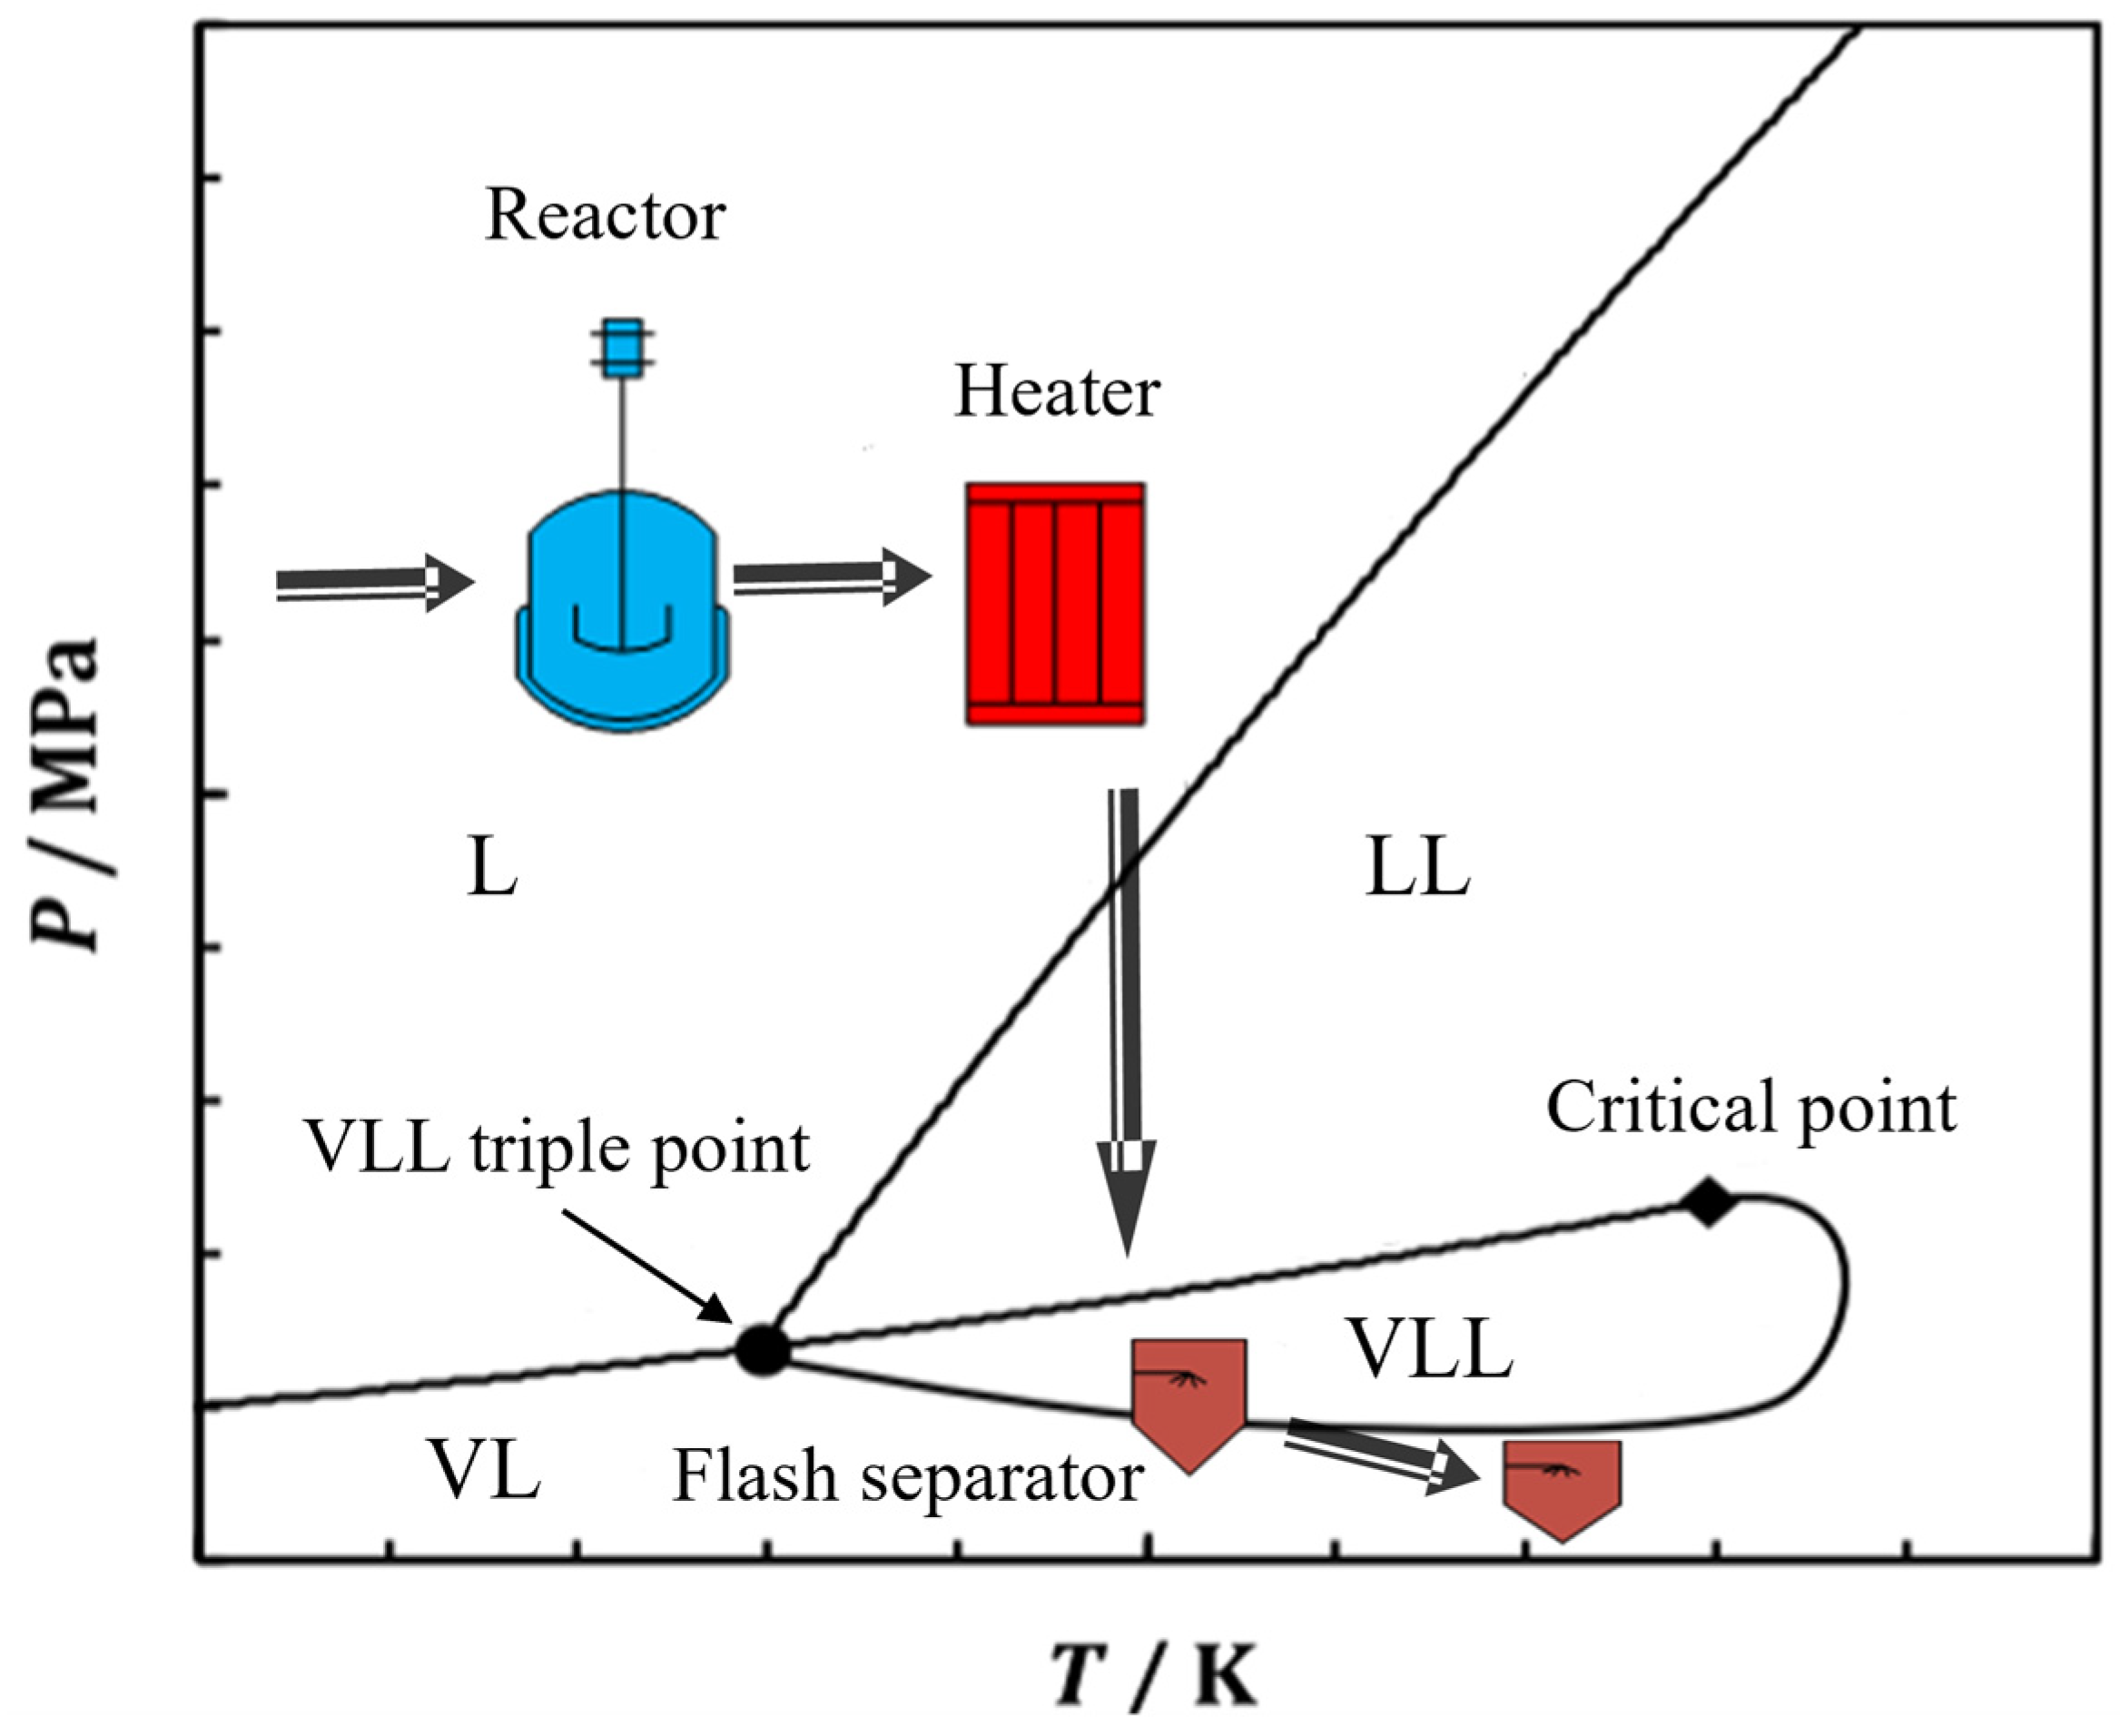
Figure 5 illustrates that, regardless of the type of α-olefin as the third component, all ternary systems exhibit LCST-type phase behavior, consistent with the binary
n-hexane + POE96k-10 system. As shown in
Figure 5a, ethylene shifts the LL line toward lower temperatures and higher pressures, effectively moving it to the upper left in the phase diagram. The liquid–liquid biphasic region expands, and as the ethylene mass fraction increases, the L → LL phase transition temperature decreases, while the transition pressure increases. The LL phase transition with 1-butene (
Figure 5b) is similar to that observed with ethylene. The addition of 1-hexene has a negligible effect on the LL phase transition (
Figure 5c). As the 1-hexene mass fraction increases from 0.0988 to 0.4982 g·g
–1, the liquid–liquid biphasic region sightly expands, with only a minor change. A similar trend is observed for the VL line. In
Figure 5d, the addition of 1-octene reduces the size of the liquid–liquid biphasic region, increases the L → LL transition temperature, and decreases the transition pressure. This effect becomes more pronounced as the 1-octene mass fraction increases.
Regarding vapor-liquid phase behavior, the addition of ethylene significantly increases the transition pressures, and 1-butene has a similar effect. Taking experimental uncertainties into account, the vapor-liquid phase transition pressure for the ethylene +
n-hexane + POE96k-10 system is consistent with the bubble point pressure measured by Nagy et al. [
44] for the ethylene +
n-hexane system, as indicated by the long-dashed line in
Figure 5a. This suggests that the polymer has a minimal impact on the L(L) → VL(L) phase transitions. 1-Octene slightly lowers the VL phase transition pressure; however, the change is less pronounced compared to the LL phase transition.
5.2.2. α-Olefin Influence on Phase Transitions
To quantitatively describe the phase transition of the α-olefin +
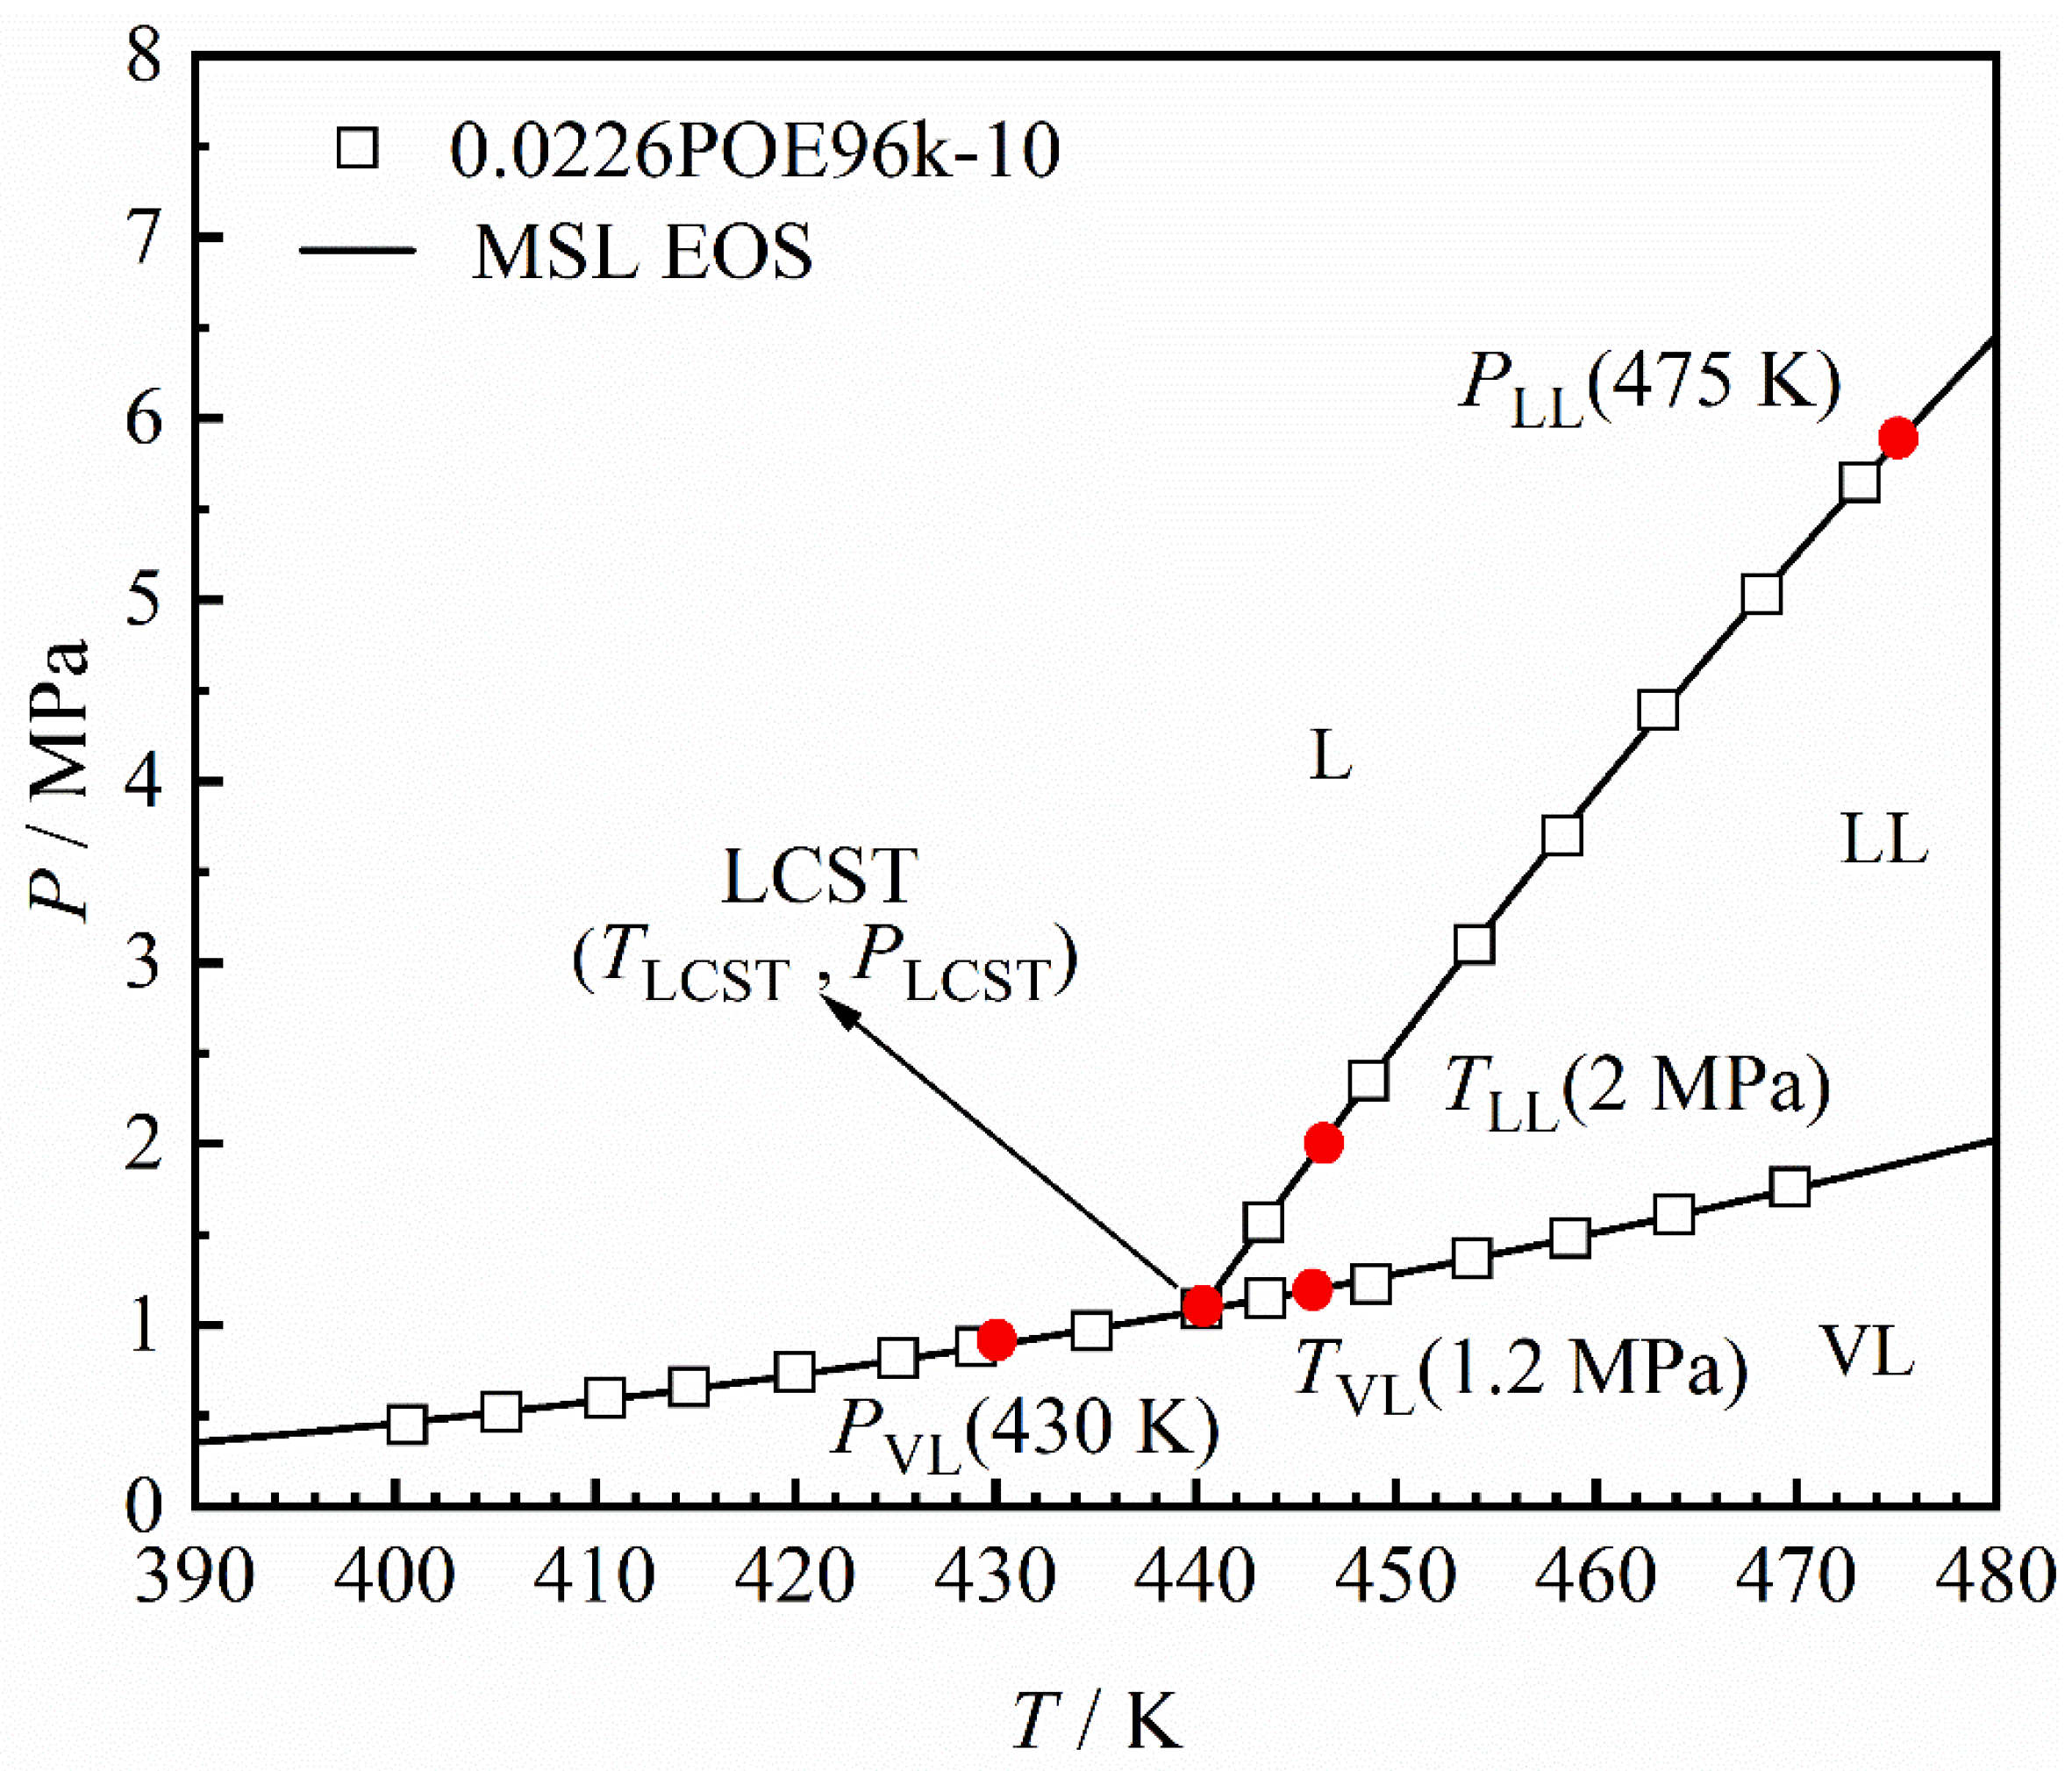
n-hexane + POE96k-10 system, five key phase transition points (corresponding to six phase transition values) were selected from the
P–
T phase diagram, as summarized in
Figure 6. These five points are the phase transition pressure on the liquid–liquid equilibrium (LLE) line at 475 K,
PLL(475 K); the phase transition temperature on the LLE line at 2 MPa,
TLL(2 MPa); the phase transition pressure on the vapor-liquid equilibrium (VLE) line at 430 K,
PVL(430 K); the phase transition temperature on the VLE line at 1.2 MPa,
TVL(1.2 MPa); and the LCST point (
TLCST,
PLCST) defined as the intersection of the LLE and VLE lines. Among these,
TLCST,
PLL(475 K), and
TLL(2 MPa) are used to analyze the liquid–liquid phase region. The selection of 475 K and 2 MPa is based on the observation that, at these conditions, different α-olefin +
n-hexane + POE96k-10 systems exhibit cloud point behavior. Alternative temperatures or pressures may also be used, provided that all systems exhibit a cloud point curve under the selected conditions. Similarly,
PLCST,
PVL(430 K), and
TVL(1.2 MPa) are used to analyze the vapor-phase behavior. The selection of 430 K and 1.2 MPa follows the same as above, ensuring that all systems undergo an L(L) → VL(L) phase transitions at these conditions.
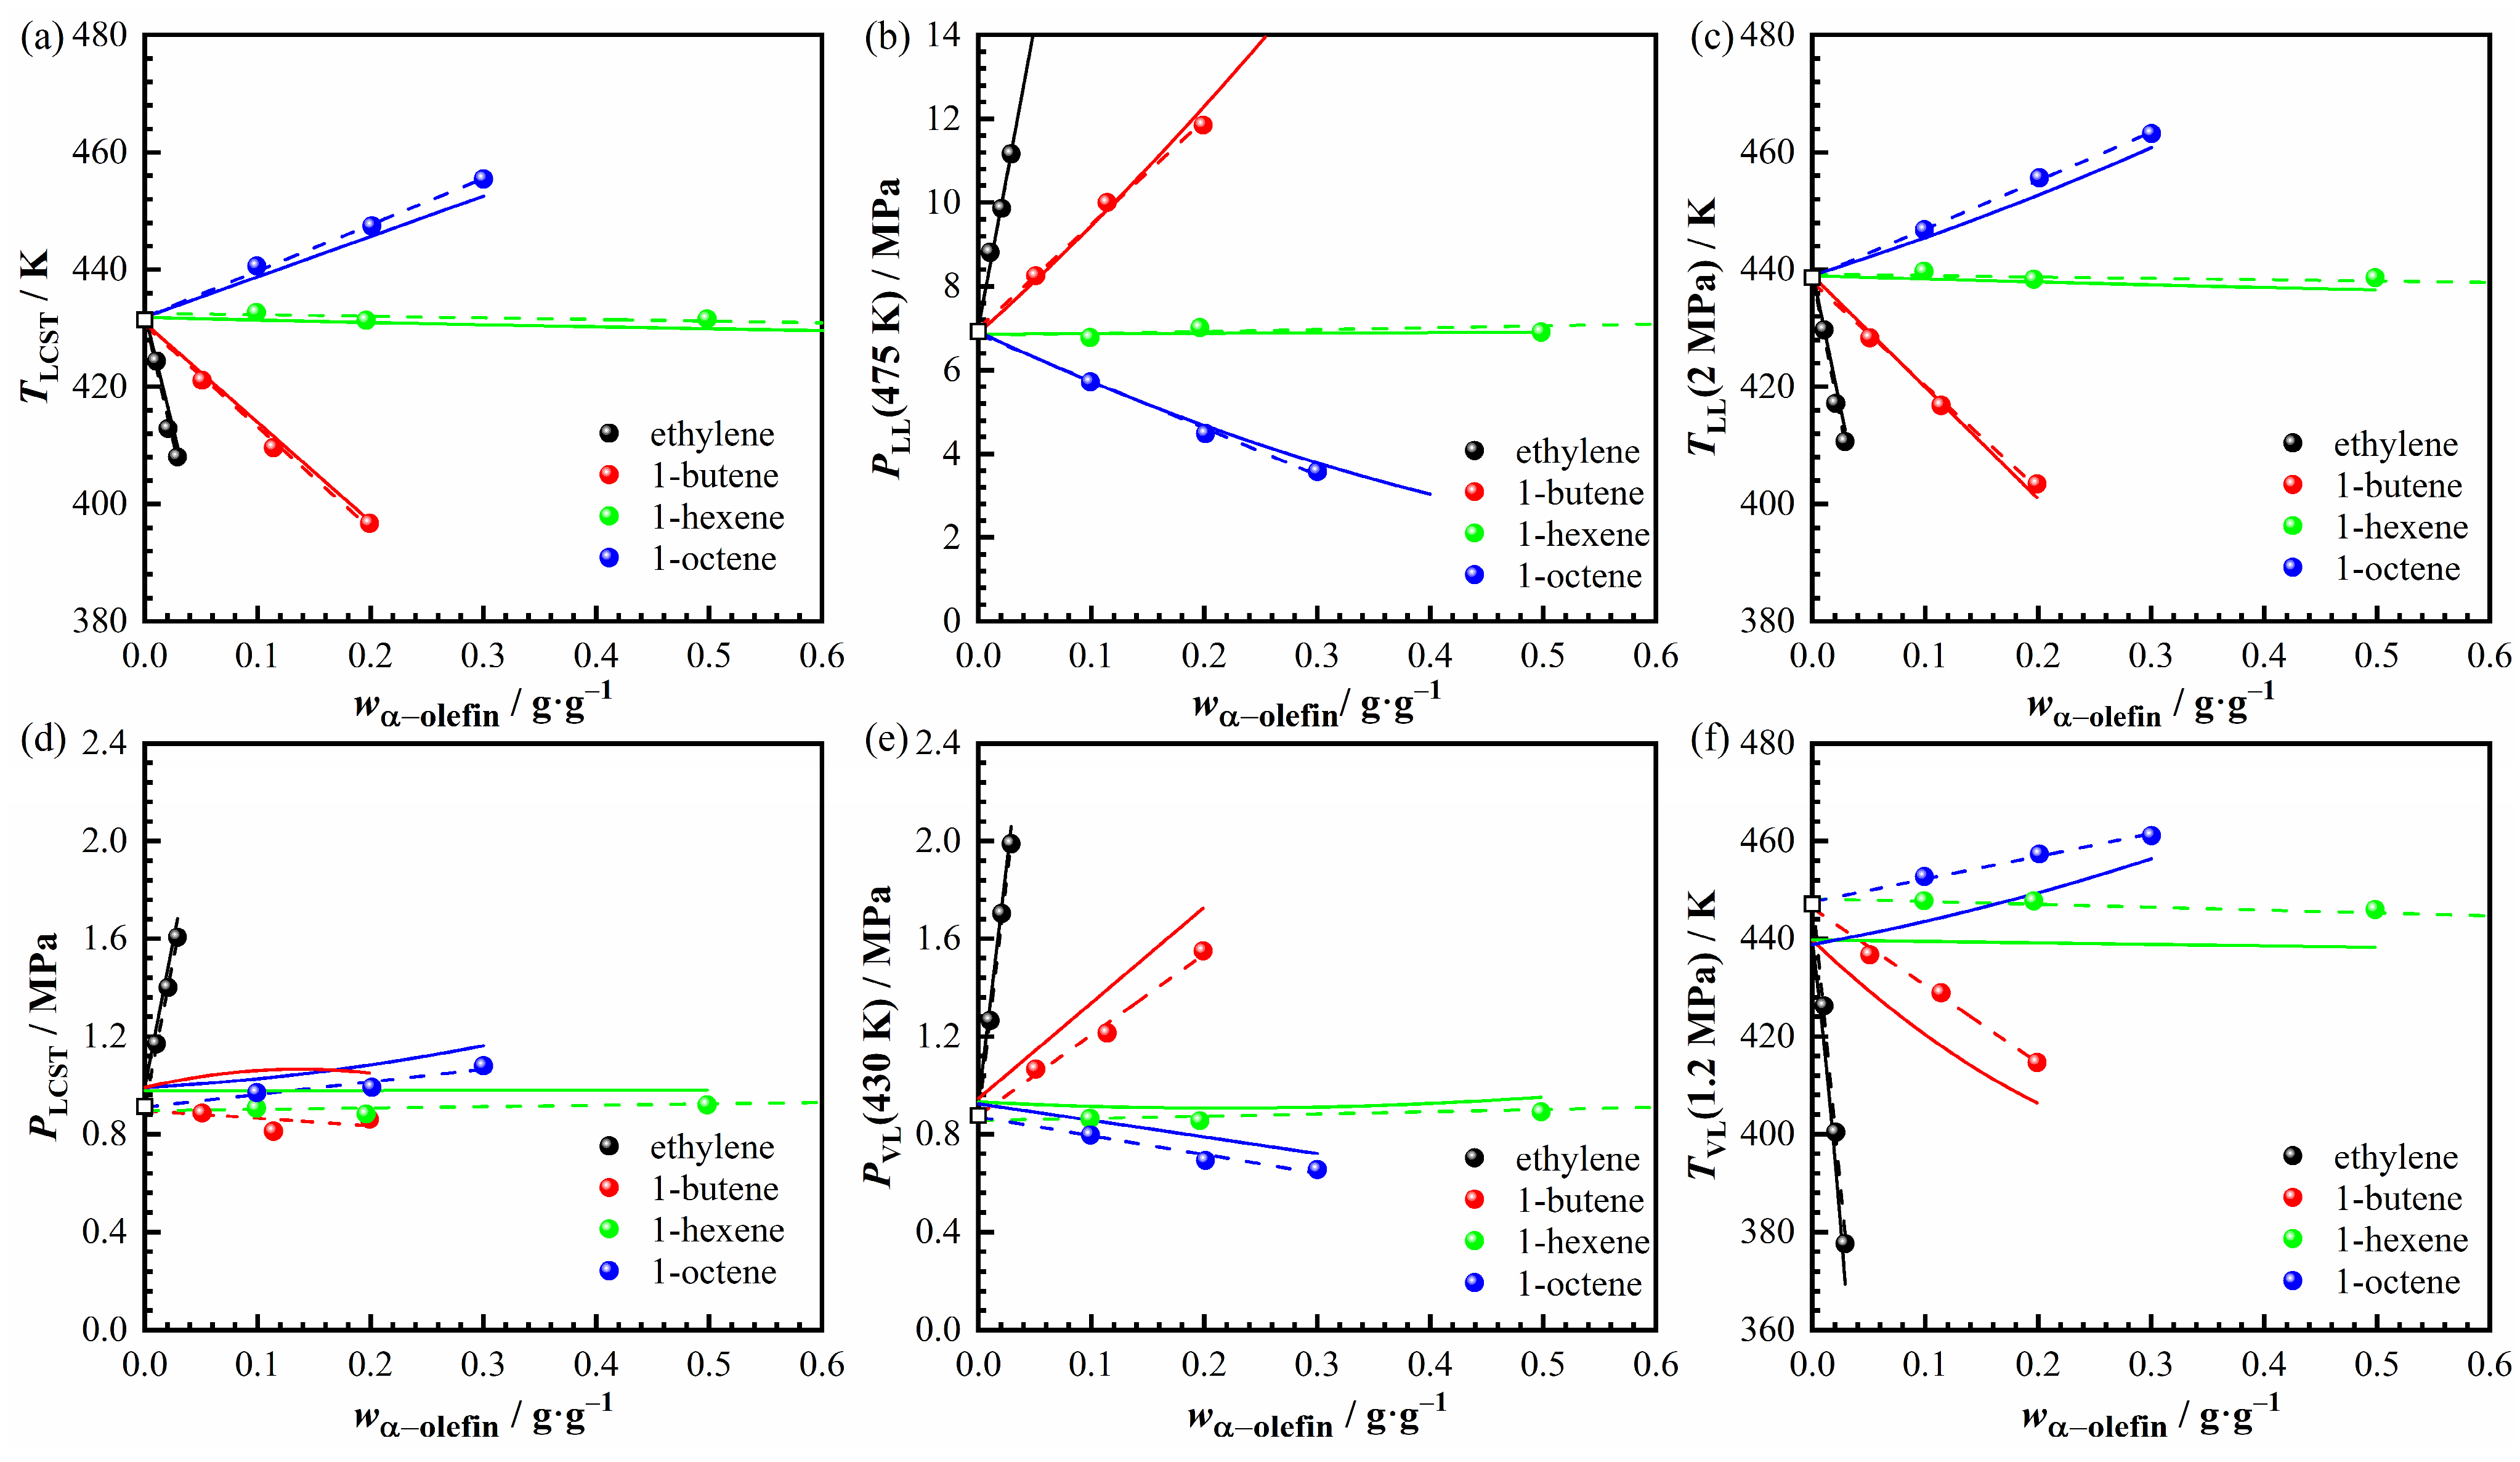
Figure 7 presents the variation in the six characteristic phase transition values described above as a function of α-olefin content in different α-olefin +
n-hexane + POE96k-10 systems. Specifically,
Figure 7a–c depict the three characteristic values related to the L → LL transition, while
Figure 7d–f represent the three characteristic values associated with L(L) → VL(L) transition. The transition temperature and pressure of L → LL or L(L) → VL(L) phase transitions generally exhibit an approximately linear trend as the α-olefin mass fraction changes. To quantify these trends, linear regressions were performed for the characteristic points and the corresponding slopes, together with the evaluation of confidence intervals. The fitted slopes and their associated 95% confidence intervals are summarized in
Table S16. Although the magnitude of the slopes varies among different α-olefins and characteristic points, the statistical analysis confirms that the observed trends are systematically resolvable beyond experimental scatter. In the nonpolar α-olefin +
n-hexane + POE system, dispersion forces dominate the intermolecular interactions [
45]. Due to their quasi-additivity nature [
46], variations in α-olefin mass fraction led to proportional changes in the intermolecular interactions between POE and the solvent (
n-hexane + α-olefin). As a result, the phase transition temperature and pressure also change proportionally as α-olefin mass fraction increases.
The linear-fit slopes in
Figure 7 are listed in
Table 8 and further illustrated in
Figure 8. These slopes represent the rates of change in phase transition temperature and pressure with respect to the α-olefin mass fraction. As shown in
Table 8, when the α-olefin is 1-hexene, the variation rates of all six indicators are close to zero, indicating that 1-hexene has little effect on phase transition temperature and pressure. This behavior can be attributed to the similar molecular chain length of 1-hexene and n-hexane, which leads to comparable dispersion interactions between POE and the two solvents. Additionally,
n-hexane and 1-hexene have nearly identical vapor pressures, resulting in a negligible influence of 1-hexene on phase behavior, which is consistent with the results in
Figure 5c. 1-Butene lowers
TLCST but has little effect on
PLCST (
Figure 7 and
Table 8). Although the decrease in
TLCST reduces the volatility of the (1-butene +
n-hexane) mixture at the LCST point, the inherently high volatility of 1-butene increases the L(L) → VL(L) phase transition pressure. These two effects offset each other, resulting in only a minor change in
PLCST.
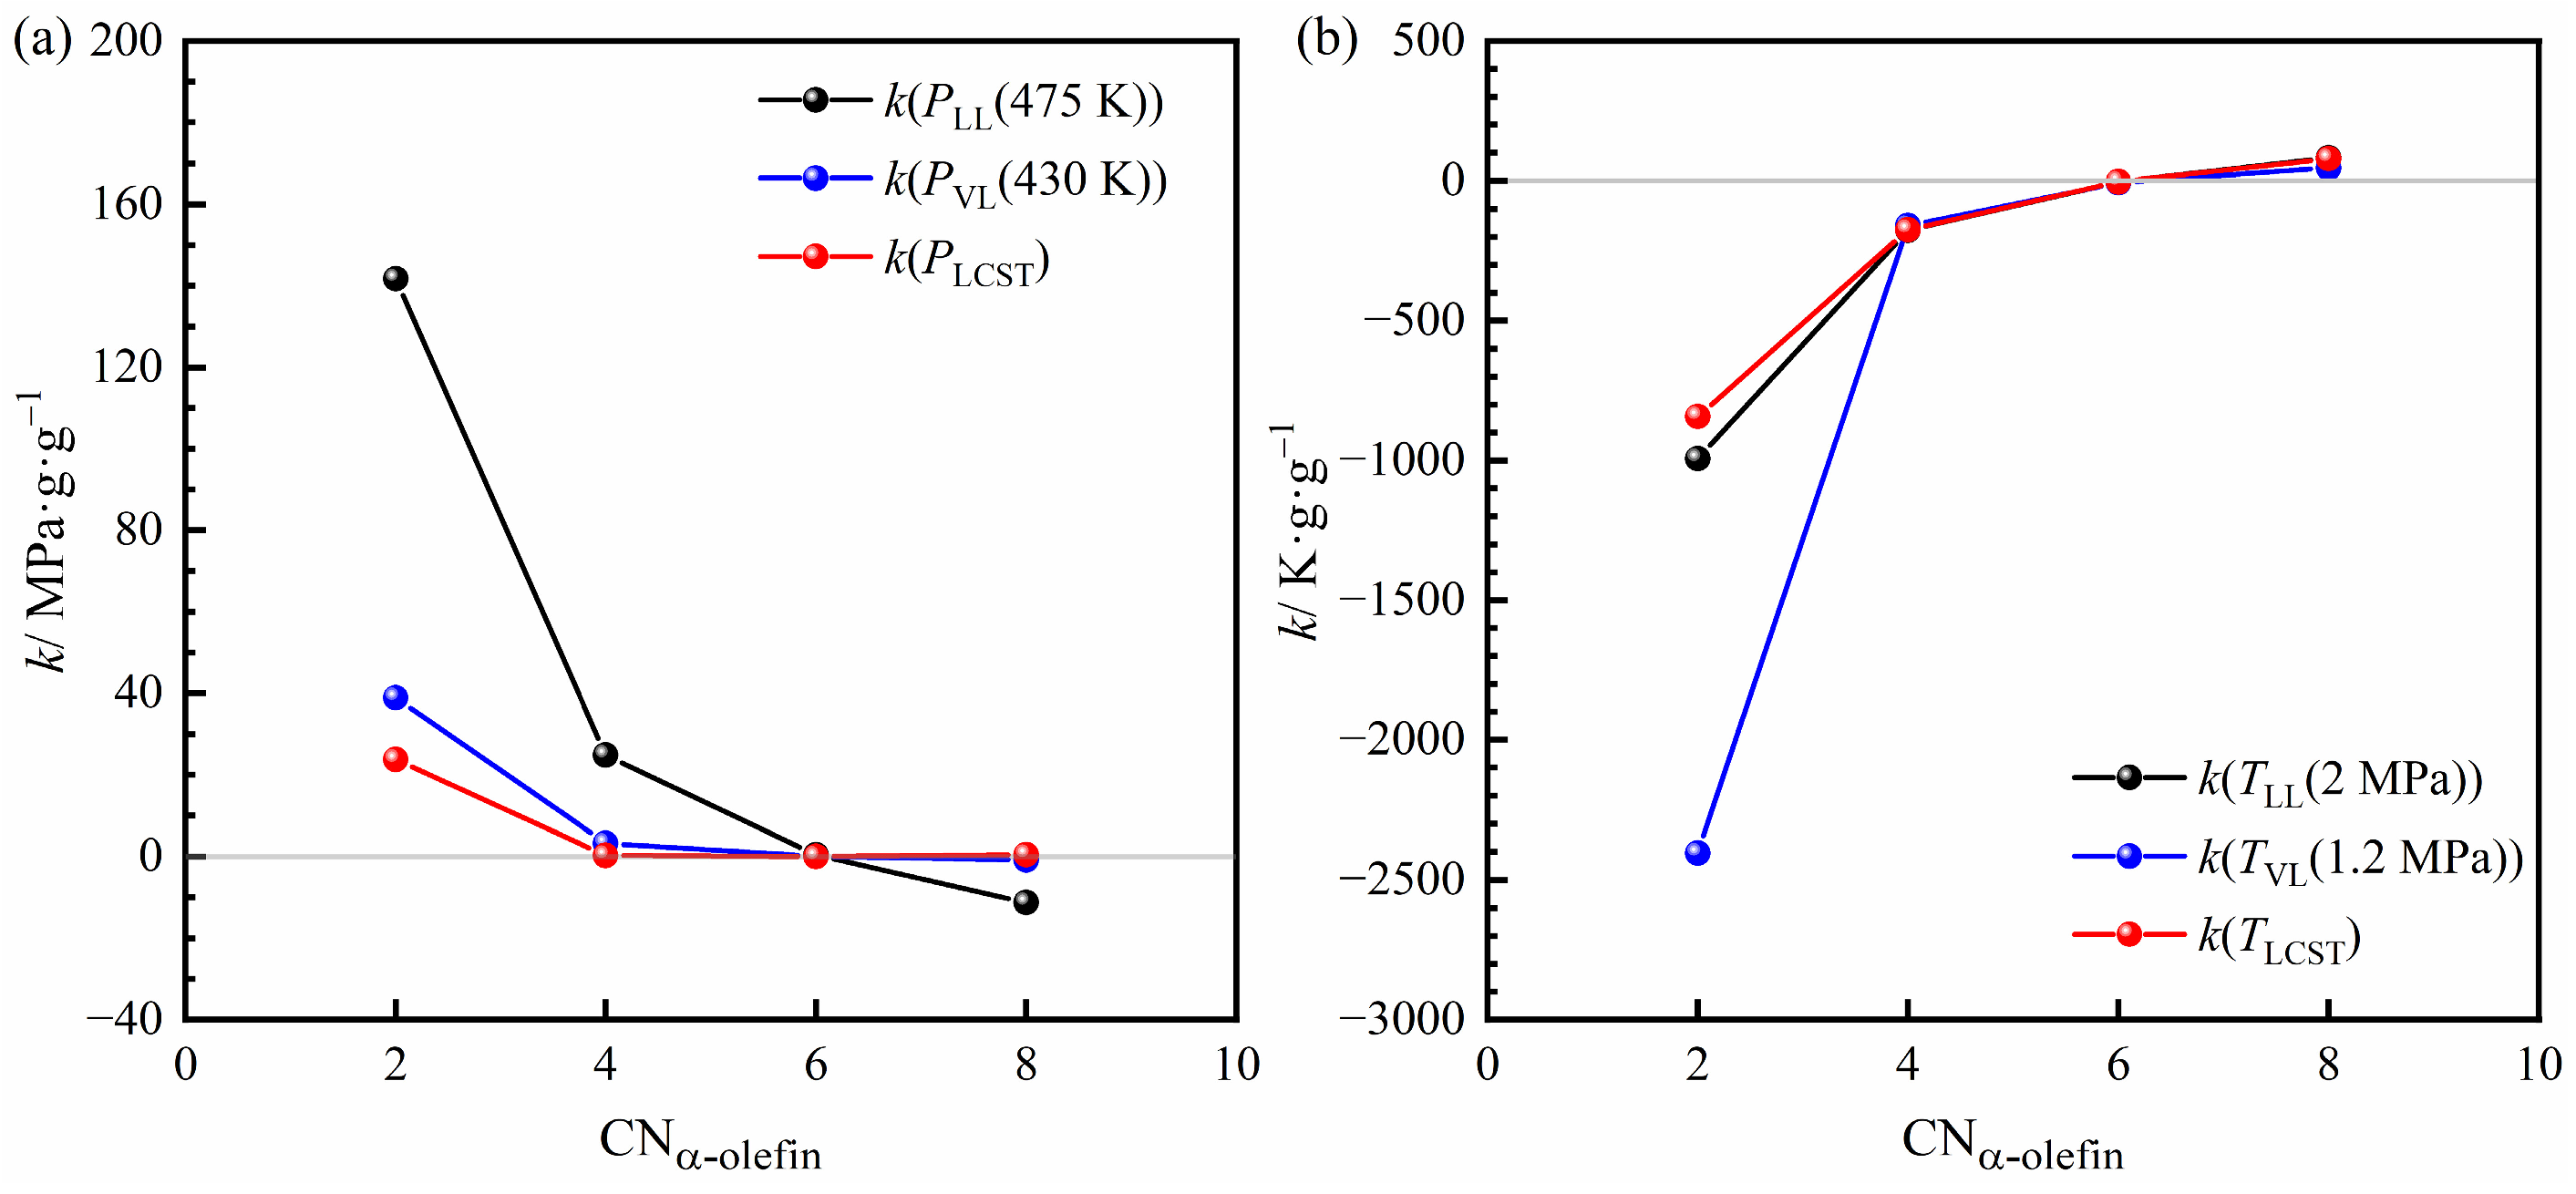
Figure 8 visually illustrates how different α-olefins influence the variation in phase transition pressure and temperature. Regarding the L → LL phase transition, the variation rates of
TLL(2 MPa) and
TLCST remain nearly identical (
Figure 8b). Meanwhile, for C
4–C
8 α-olefins, the variation rate of
PLCST remains close to zero. The variation rate of
PLL(475 K) shifts from positive for 1-butene to nearly zero for 1-hexene and then to negative for 1-octene. This means
PLL(475 K) increases with 1-butene, remains unchanged with 1-hexene, and decreases with 1-octene. These changes in
PLL(475 K) are primarily influenced by the corresponding shifts in phase transition temperature: the decrease for 1-butene and the increase for 1-octene. Thus, for the L → LL phase transition, C
4–C
8 α-olefins mainly affect phase transition temperature rather than pressure.
The addition of 1-butene lowers the L → LL phase transition temperature, while 1-hexene has little effect. In contrast, 1-octene increases the phase transition temperature (
Figure 5 and
Figure 8). Since α-olefins share a similar chemical structure with
n-alkanes, their intermolecular interactions resemble those of
n-alkanes with the same carbon number. Moreover, the dispersion force involving the polymer and α-olefin generally increases with the molar mass of the α-olefin. As a result, the dispersion force involving the polymer and 1-butene is weaker than that with
n-hexane, whereas for 1-hexene, it remains similar to that with
n-hexane. In contrast, the dispersion force between the polymer and 1-octene is stronger. This phenomenon can be explained by the thermodynamic criterion for homogeneous solution formation from multiple pure components:
According to this equation, temperature correlates with Δ
Hm/Δ
Sm. Since phase separation occurs upon heating in LCST-type phase behavior, Δ
Sm < 0 [
47]. Stronger dispersion forces make mixing more exothermic, leading to a more negative Δ
Hm and, consequently, a higher phase transition temperature. Conversely, weaker dispersion forces between the polymer and solvent result in a lower L → LL transition temperature.
As shown in
Figure 8, when the α-olefin is 1-hexene or 1-octene, their effects on the VL phase transition pressure are nearly identical. Although the vapor pressures of 1-hexene and 1-octene differ, the VL phase transition is still predominantly governed by n-hexane. In contrast, α-olefins with higher volatility than n-hexane, such as ethylene or 1-butene, exert a significant influence on the VL phase transition pressure. This influence becomes more pronounced as the α-olefin carbon number decreases.
Compared to other α-olefins, ethylene exhibits the most pronounced influence on both phase-transition temperature and pressure. While dispersion interactions generally increase with the molar mass of α-olefins, ethylene shows a clear deviation from this trend and behaves as a strong anti-solvent. This behavior cannot be explained by dispersion forces alone and suggests a reduced effective affinity between ethylene and the polymer–solvent system. Unlike
n-alkanes, α-olefins contain a carbon–carbon double bond, which may introduce additional electronic effects. For ethylene, owing to its short molecular chain, such effects may become relatively more significant compared to dispersive interactions. From a thermodynamic perspective, this behavior can be qualitatively rationalized in terms of a weaker effective polymer–solvent affinity, which reduces polymer solvation and shifts the L → LL phase boundary to lower temperatures. In the absence of direct quantitative thermodynamic analysis, this interpretation should be regarded as a plausible qualitative explanation rather than a definitive mechanism. Additionally, ethylene significantly increases
PLCST and
PVL(430 K) in
Figure 8a. This is because ethylene has much higher volatility than
n-hexane, making the L(L) → VL(L) phase behavior primarily controlled by ethylene. Ethylene simultaneously reduces the transition temperature and markedly raises the transition pressure.





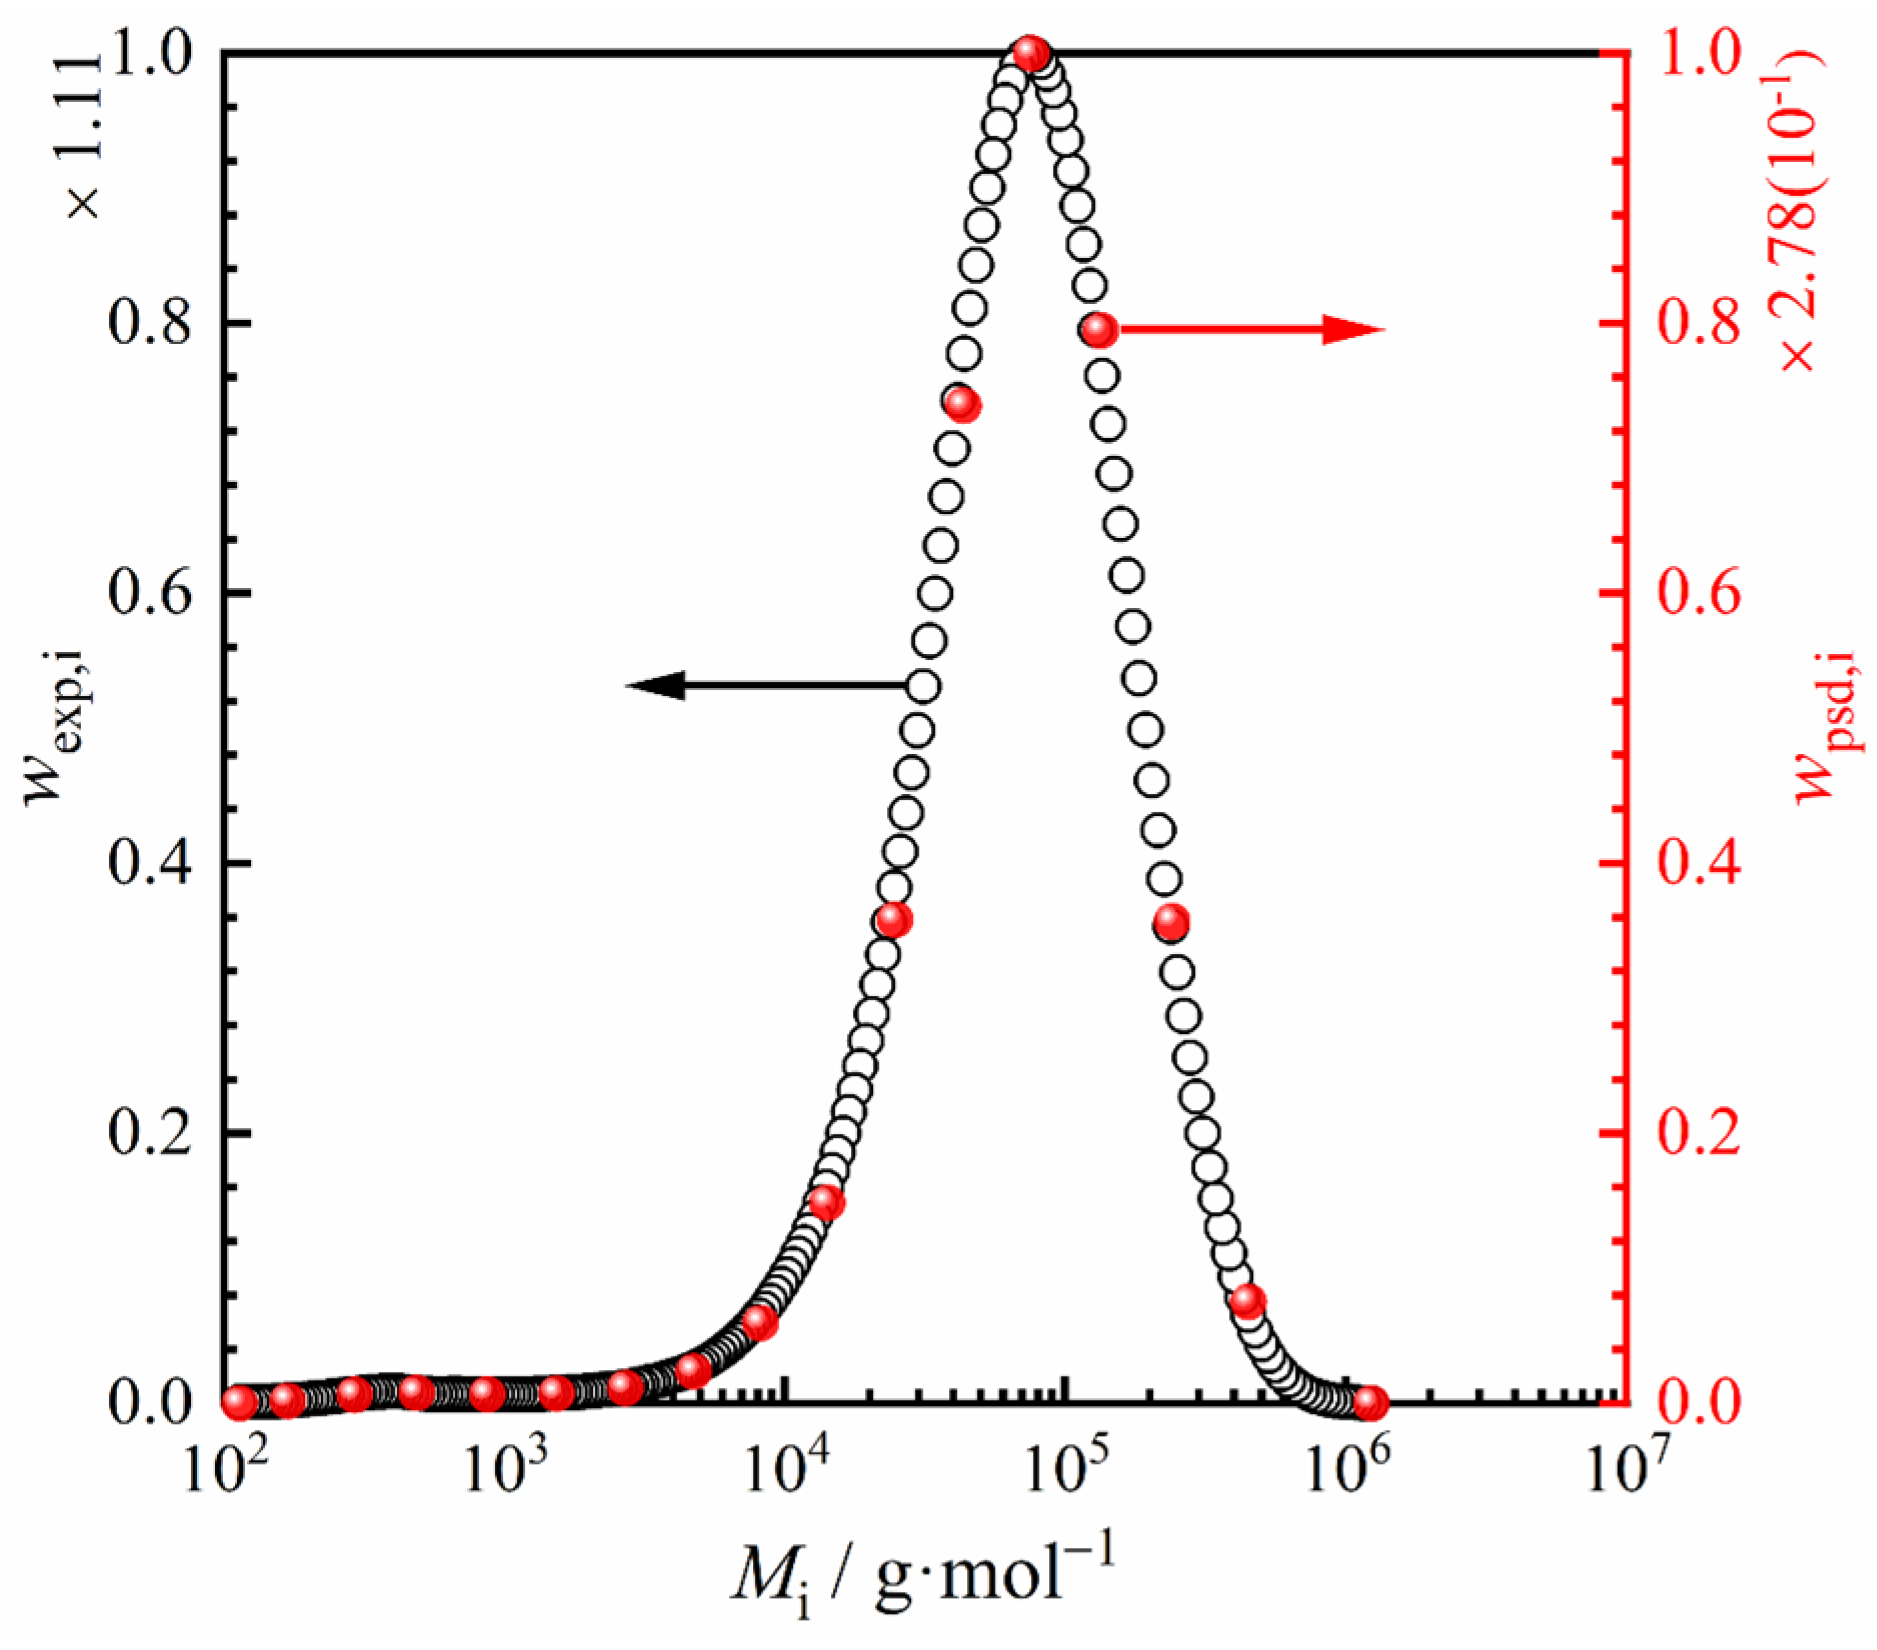
 ) pseudocomponents from Table 3.