Article

Response of Soil Microbial Communities to Oil Pollution and Remediation in the Yellow River Delta

1
Sinopec Petroleum Engineering Corporation, Dongying 257000, China
2
Shandong Key Laboratory of Eco-Environmental Science for the Yellow River Delta, Shandong University of Aeronautics, Binzhou 256600, China
*
Author to whom correspondence should be addressed.
Diversity 2026, 18(4), 206; https://doi.org/10.3390/d18040206
Submission received: 25 January 2026 / Revised: 25 February 2026 / Accepted: 25 February 2026 / Published: 31 March 2026
(This article belongs to the Section Microbial Diversity and Culture Collections)

Abstract

Soil microorganisms, as a crucial component of the soil ecosystem, play an essential role in maintaining soil health, promoting nutrient cycling, and ensuring ecological stability. Nevertheless, with the rapid progress of industrialization, crude oil pollution has emerged as a significant environmental hazard, notably affecting the structure and functionality of soil microbial communities. The Yellow River Delta, a vital wetland ecosystem and a key oil-producing area in China, has been exposed to crude oil contamination for a long time, leading to a substantial decline in its ecological functions. This study elucidates the effects of crude oil pollution on soil microorganisms in the Yellow River Delta by comparing the microbial community composition between long-term contaminated sites and sites that have undergone phytoremediation. By comprehensively analyzing the ecological responses and remediation potential of microbial communities, this research aims to offer a scientific basis for regional ecological restoration and sustainable development.

1. Introduction

Soil microbial health refers to the overall condition in which the microbial community maintains its diversity, ecological functionality, and capacity for environmental adaptation [1,2,3]. It serves as a key indicator for assessing the stability and productivity of the soil ecosystem. Microorganisms play a crucial role in the biogeochemical cycling of elements such as carbon, nitrogen, and phosphorus, transforming atmospheric carbon dioxide and nitrogen into forms that plants can absorb and utilize [4,5]. By decomposing plant and animal residues as well as organic pollutants, microorganisms facilitate the synthesis and breakdown of soil organic matter—a process essential for the development of soil fertility. Studies indicate that microbial activity contributes to the formation of soil aggregates larger than 0.25 mm [4]. These aggregates function like sponges, retaining water and air while sequestering nutrients. A healthy soil microbial community is characterized by high diversity and stability, enabling it to effectively respond to environmental changes. However, industrial pollutants such as crude oil exert significant and direct toxic effects on soil microorganisms, potentially disrupting community structure and impairing functional capabilities [6]. The Yellow River Delta represents a significant wetland ecosystem and is China’s key oil production base. The primary sources of pollution include oil leakage incidents during extraction and transportation processes, as well as residual contamination from historically abandoned oil wells. Studies indicate that surface sediments near sealed and abandoned wells are contaminated with polycyclic aromatic hydrocarbons (PAHs), with varying ecological recovery observed across sites closed at different times [7]. These differences highlight the influence of pollution duration on ecosystem restoration [8]. The pollution level is relatively severe in concentrated oil extraction areas, such as the Binan Oilfield, whereas it is comparatively mild in the core zones of nature reserves. This spatial distribution highlights the necessity of formulating region-specific ecological risk assessment and remediation strategies. Persistent organic pollutants, particularly polycyclic aromatic hydrocarbons (PAHs) present in crude oil, exhibit “carcinogenic, teratogenic, and mutagenic” properties, posing threats not only to soil ecosystem health but also potentially impacting human health through the food chain [9]. It is important to note, however, that not all constituents of crude oil uniformly exhibit these toxicological effects, as toxicity varies significantly depending on the specific compound structure, concentration, and environmental bioavailability.
Phytoremediation is an environmentally friendly technique that employs plants and their rhizosphere microbial communities to eliminate soil contaminants [10,11]. Previous studies have indicated that microbial diversity in crude oil-contaminated soil typically declines by over 40%, with sensitive microbial groups, such as nitrogen-fixing bacteria, experiencing up to 90% reductions [12]. The soil microbial community undergoes adaptive structural and functional changes throughout the remediation process, which indicate remediation efficacy and actively influence the overall remediation dynamics [12,13,14]. Microbial remediation, which enhances natural processes, offers several advantages, including low cost, environmental sustainability, in situ treatment, and the absence of secondary pollution [15,16]. Research on microbial remediation of crude oil-contaminated soil in the Yellow River Delta holds theoretical and practical significance. Theoretically, this research contributes to a deeper understanding of microbial adaptation mechanisms and community succession patterns under extreme environmental stress. The findings can directly inform regional ecological restoration efforts and provide technical support for balancing petroleum resource development and environmental protection. Specifically, this study delves into the mechanisms by which crude oil pollution impacts soil microorganisms in the Yellow River Delta, as well as the changes in microbial community composition prior to and subsequent to the application of phytoremediation technology. By comprehensively analyzing the ecological responses and remediation potential of microbial communities, this research aims to offer a scientific basis for regional ecological restoration and sustainable development.

2. Materials and Methods

2.1. Sample Collection

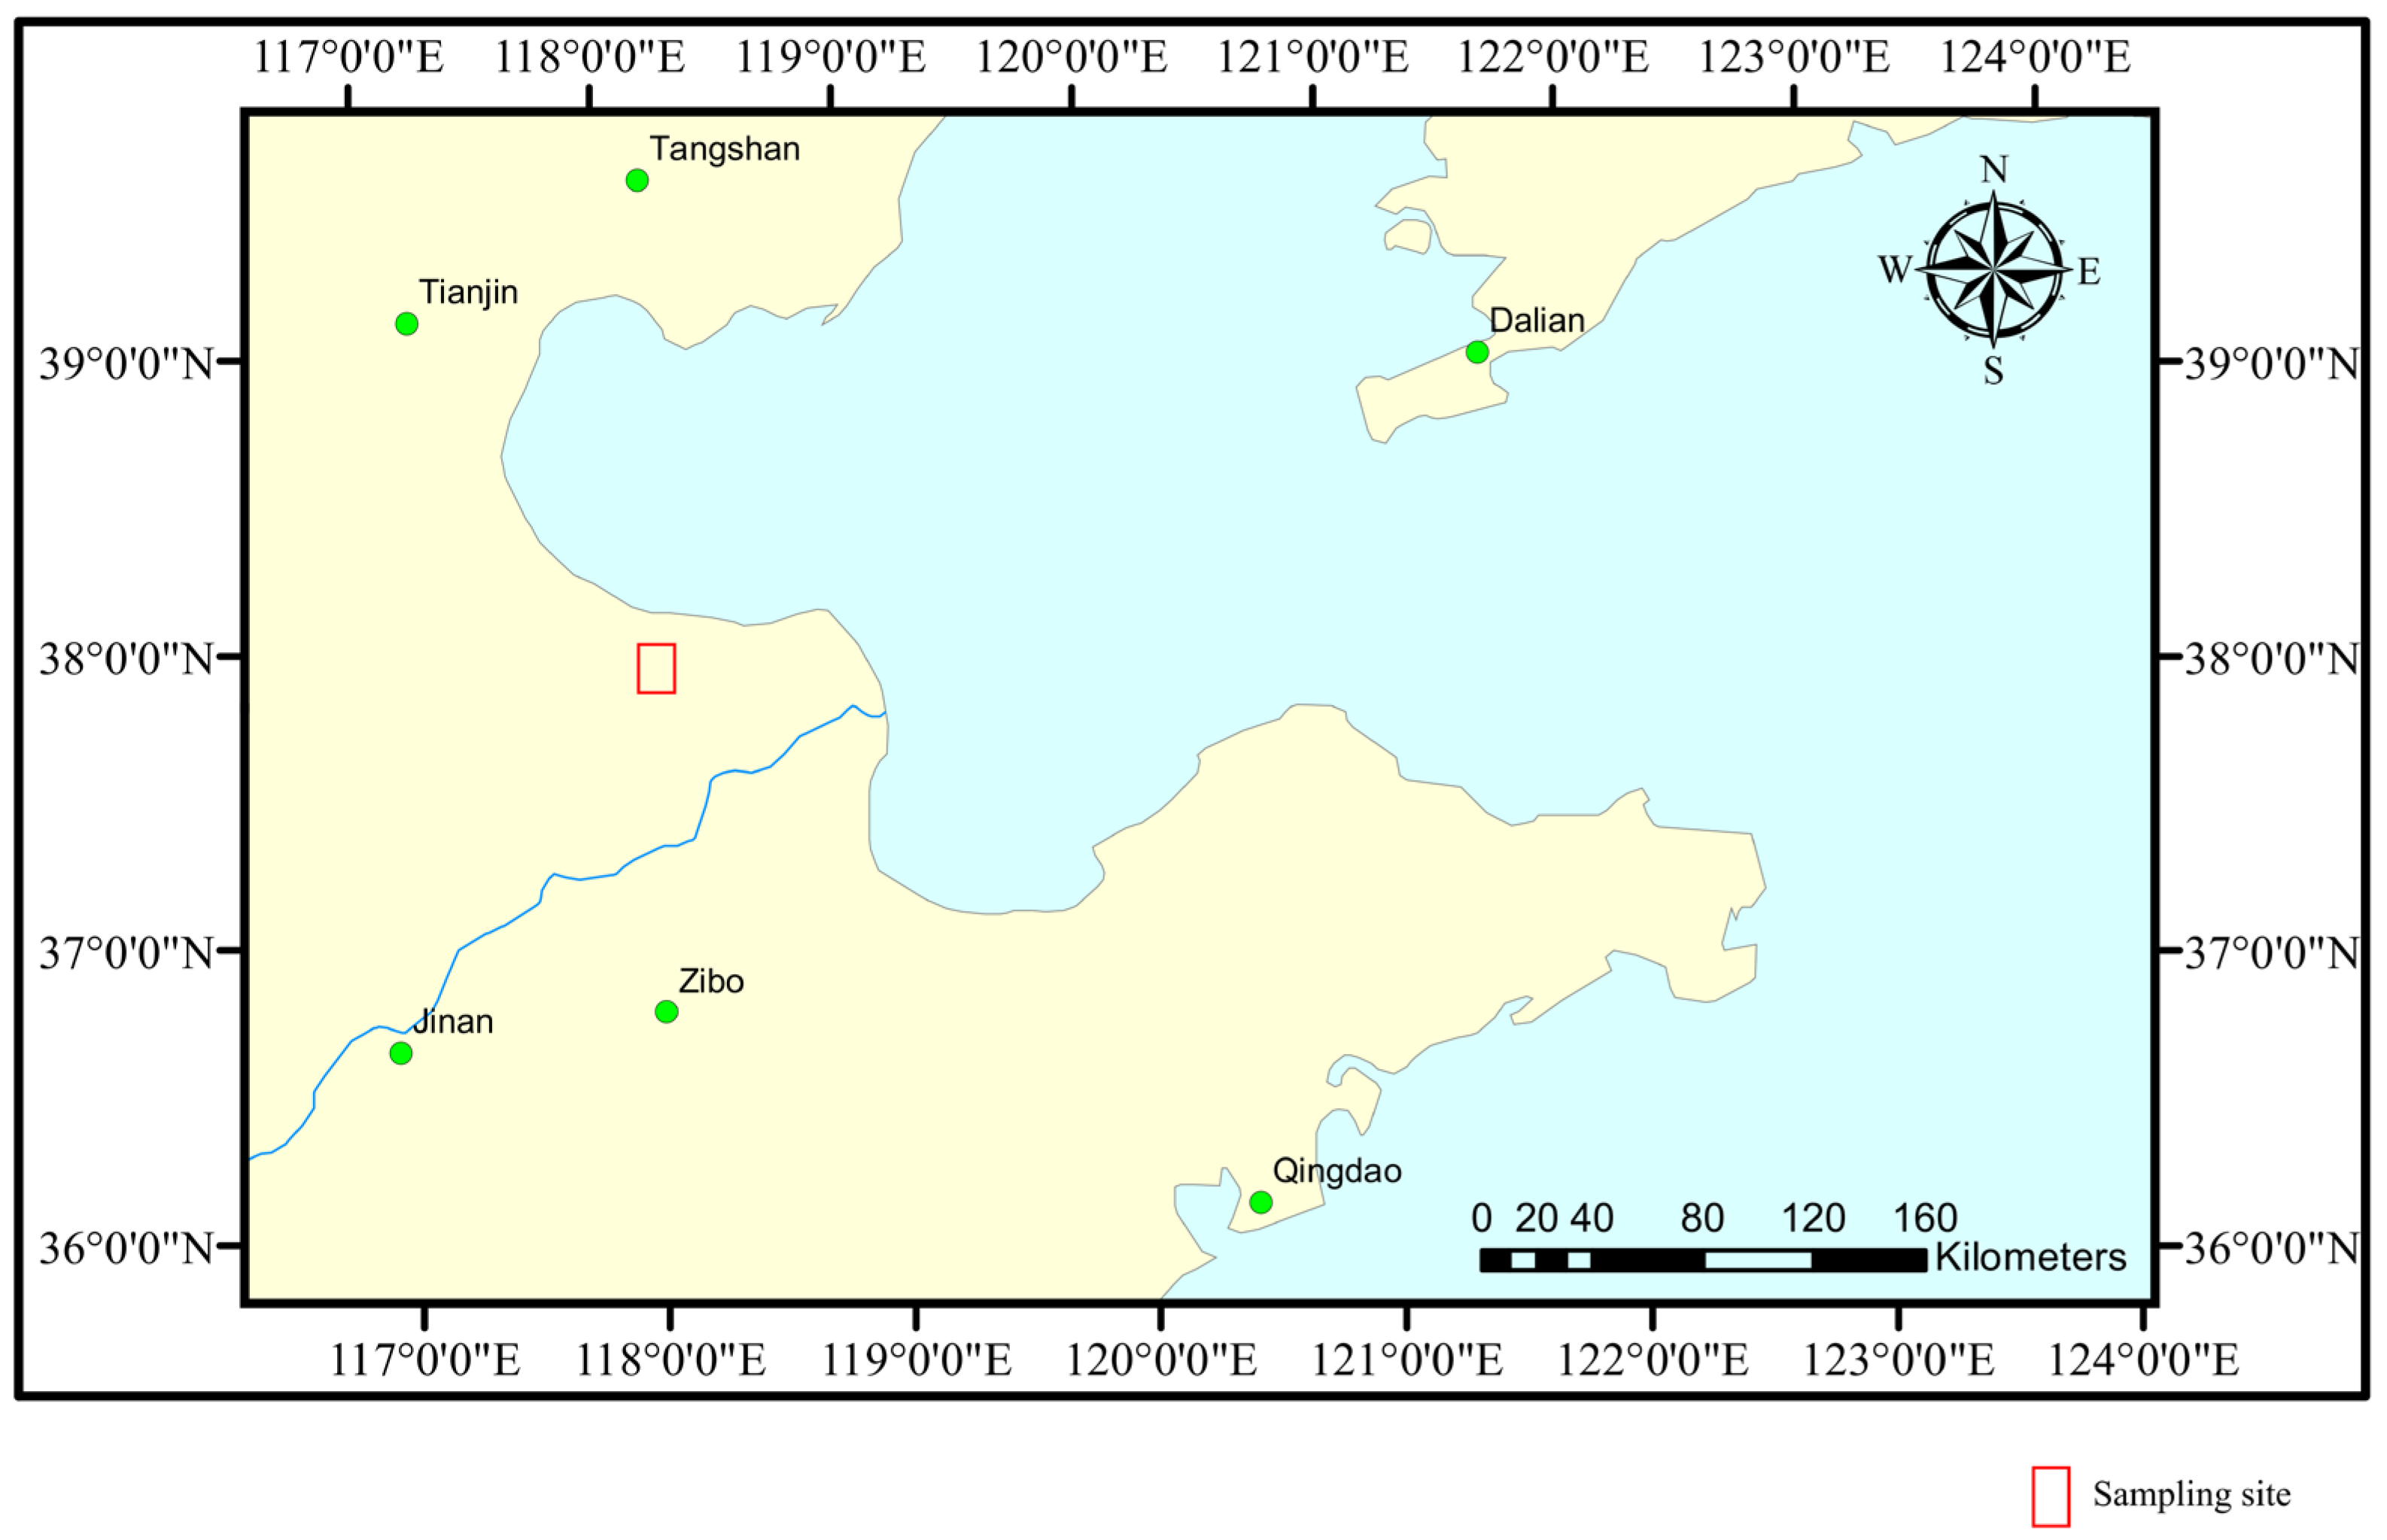
For the purposes of this research, a total of 30 soil samples were systematically collected (Figure 1). Specifically, 15 samples were collected from a petroleum-contaminated site (Group S, S1–S15) and 15 samples from a phytoremediated site (Group R, R1–R15). At each site, three 10 m × 10 m replicate plots were established. Within each plot, five sampling points were randomly selected, resulting in five independent soil samples per plot. At the bioremediated site, the grass Cynodon dactylon was planted as the primary remediation plant, along with scattered herbaceous species. Both sites share similar pedological conditions, and the soil is classified as coastal saline–alkaline fluvo-aquic soil (Calcaric Fluvisol, FAO classification). The region has a warm–temperate monsoon climate, with a mean annual temperature of 12.6 °C and a mean annual precipitation of 560 mm, which is mainly concentrated in the summer months. The salt content increases with depth and during the dry seasons. Soil physicochemical properties (e.g., pH, electrical conductivity, total organic carbon, total nitrogen, total petroleum hydrocarbons, and polycyclic aromatic hydrocarbons) were not measured in this study. Therefore, direct correlations between microbial community shifts and specific environmental variables could not be established. At both sites, after removing surface litter, bulk soil samples were collected from the upper 0–20 cm layer. To avoid rhizosphere effects and ensure that the samples represent the overall soil microbial community rather than root-associated assemblages, at the remediated site, samples were taken at least 30 cm away from any plant stem. Within each site, a stratified random sampling design was adopted. All samples were collected using sterile spatulas and placed into sterile polyethylene bags.

2.2. DNA Extraction, PCR Amplification, and Sequencing

Total genomic DNA was extracted from 0.5 g of each soil sample using the DNeasy PowerSoil Pro Kit (Qiagen, Hilden, Germany) according to the manufacturer’s protocol [17]. Cell lysis was enhanced by bead-beating using a FastPrep-24™ 5G instrument (MP Biomedicals, Santa Ana, CA, USA). An extraction blank control was included to monitor potential contamination during the extraction procedure. The V3–V4 hypervariable region of the bacterial 16S rRNA gene was amplified via PCR using the following primers: forward primer 5′-ACTCCTACGGGAGGCAGCA-3′ and reverse primer 5′-GGACTACHVGGGTWTCTAAT-3′ [18], with Illumina sequencing adapters appended to both ends. PCR amplification was performed in a T100 Thermal Cycler (Bio-Rad, Hercules, CA, USA) under the following conditions: initial denaturation at 95 °C for 3 min, followed by 30 cycles of denaturation at 95 °C for 30 s, annealing at 55 °C for 30 s, and extension at 72 °C for 30 s, with a final extension at 72 °C for 5 min [19]. Each PCR reaction included negative controls (sterile water instead of DNA template) to verify the absence of contamination. PCR products were purified using magnetic bead-based cleanup kits (Agencourt AMPure XP, Beckman Coulter, Brea, CA, USA), quantified fluorometrically using the Qubit™ dsDNA HS Assay Kit (Invitrogen, Carlsbad, CA, USA), and normalized to equimolar concentrations prior to library construction [19]. Libraries meeting quality control criteria were sequenced on an Illumina NovaSeq 6000 platform with paired-end 250 bp (PE250) chemistry. No mock community was included in the sequencing run.

2.3. Data Processing and Bioinformatics Analysis

Raw reads were subjected to processing using Trimmomatic v 0.39 for adapter trimming and quality filtering (with a Phred score of ≥20 and a minimum length of ≥150 bp). Subsequently, primers were removed using cutadapt. High-quality sequences were then processed by applying the DADA2 v1.16 (via QIIME2 v2021.4) algorithm. This algorithm was used for error correction, sequence denoising, merging of paired-end reads, and chimera detection and removal, ultimately yielding high-resolution Amplicon Sequence Variants (ASVs). ASVs were employed as operational units in lieu of traditional OTUs because of their enhanced resolution at the single-nucleotide level, which allows for more accurate taxonomic discrimination. Representative ASV sequences were classified across taxonomic ranks, ranging from kingdom to species, using the naïve Bayes classifier implemented in QIIME2. This classifier was trained against the SILVA 138 reference database (release 138). Alpha diversity metrics, such as Chao1 and ACE (which are estimators of species richness), and Shannon and Simpson indices (which are measures of diversity), were calculated based on rarefied data. To address the issue of uneven sequencing depth across samples, all samples were rarefied to 30,000 sequences per sample, which was equivalent to the minimum sequence count observed after quality filtering.
Rarefaction curves and rank-abundance distributions were generated to comprehensively assess the sequencing depth and community evenness. The differences in the composition of microbial communities between groups were precisely evaluated using Bray–Curtis, Jaccard, weighted, and unweighted UniFrac distance matrices. Principal coordinate analysis (PCoA) and non-metric multidimensional scaling (NMDS) were carried out to vividly visualize the intergroup dissimilarities, which were further supplemented by UPGMA hierarchical clustering. The statistical significance of group separation was rigorously tested using PERMANOVA (Adonis test, 999 permutations) and ANOSIM (Analysis of Similarities). LEfSe (Linear Discriminant Analysis Effect Size, v1.0) was applied to accurately identify the taxa that were significantly enriched in either group, with an LDA threshold > 4.0 and a p-value < 0.05. The results were further validated using Metastats and one-way ANOVA with a post hoc Tukey’s HSD test when appropriate. The functional profiles of the microbial communities were inferred using PICRUSt2 v2.4.1 based on ASV phylogeny, which was used to predict the abundances of KEGG orthologs and COG categories. Additionally, FAPROTAX v1.2.4 was employed to predict the potential ecological functions associated with biogeochemical cycling. For functional pathway comparisons between groups, pairwise t-tests were performed among different groups, with a p-value threshold of 0.05.

3. Results

3.1. Overview of Sequencing Data and Quality Assessment

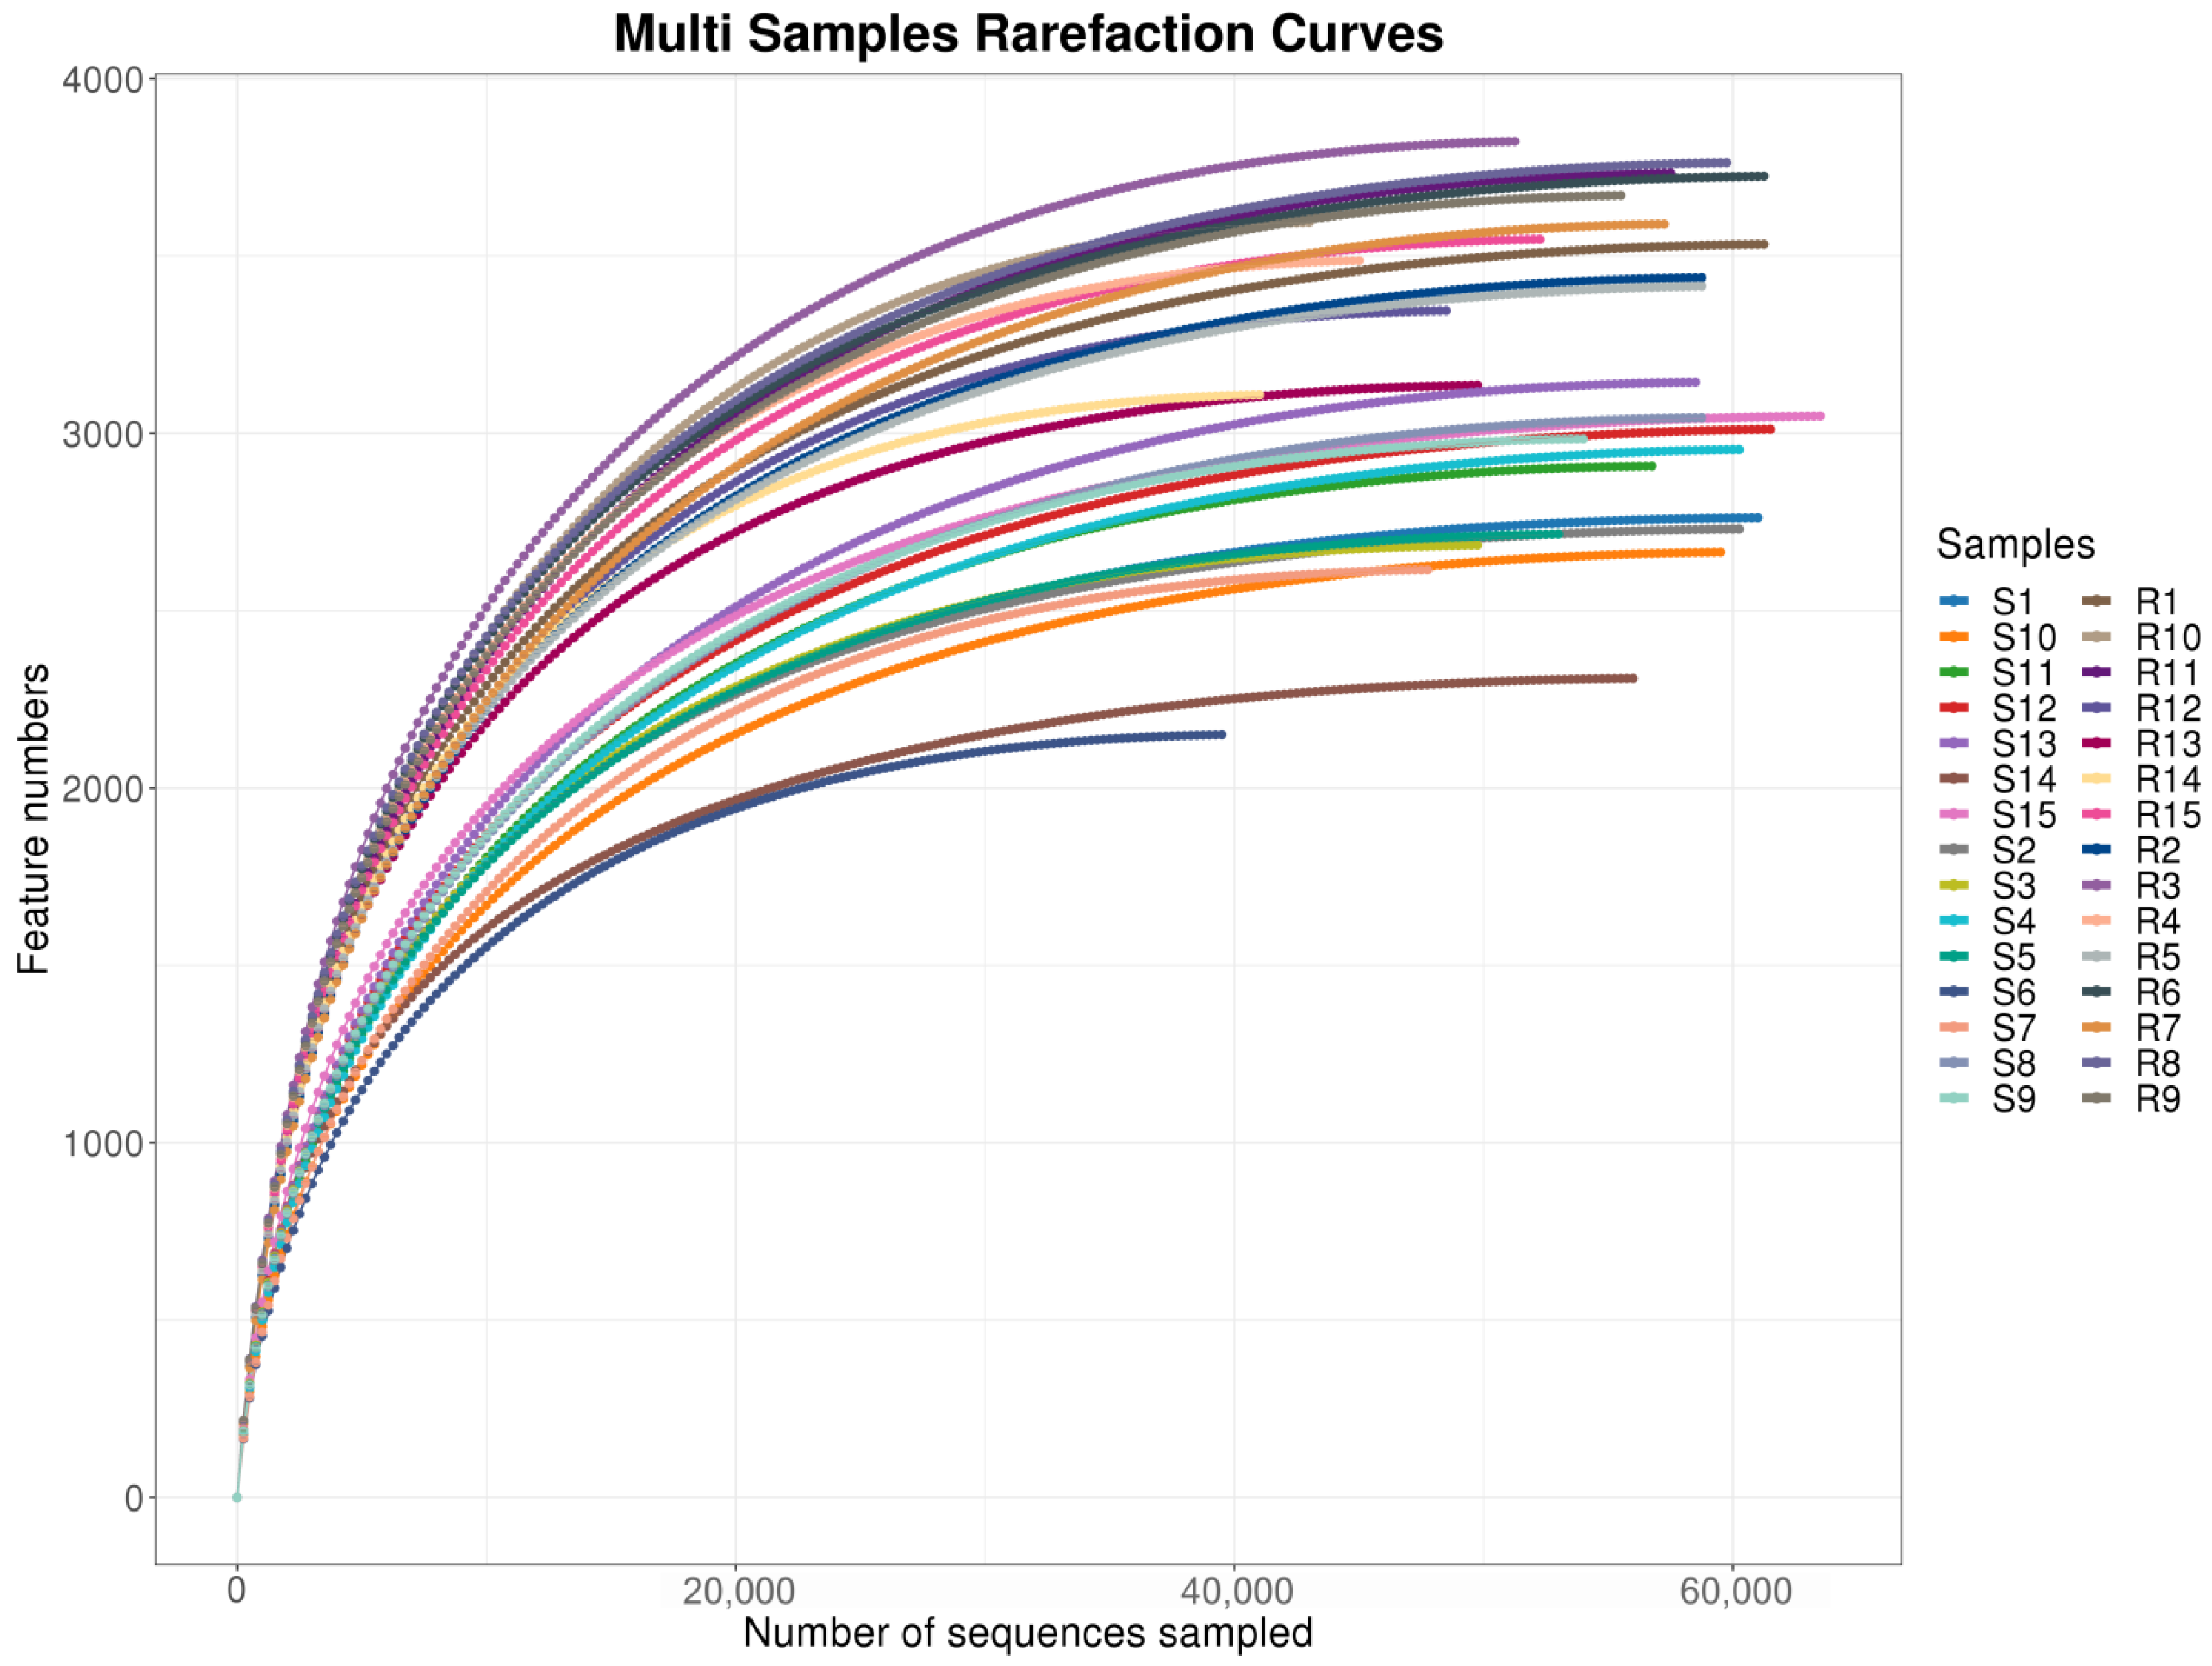
A total of 2,204,712 paired-end raw reads were generated from 30 samples (Figure 2). Following rigorous quality filtering, sequence assembly, and removal of chimeric sequences, 1,644,405 high-quality, non-chimeric reads were retained, with an average of approximately 54,813 reads per sample. Per-sample read counts ranged from 40,247 (sample S6) to 64,264 (sample S15), ensuring sufficient sequencing depth for all samples. For alpha and beta diversity analyses, all samples were rarefied to 30,000 sequences per sample to account for uneven sequencing depth. The majority of the processed sequences ranged between 400 and 450 base pairs in length. The coverage index for all samples exceeded 0.998, indicating that the sequencing depth adequately captured the vast majority of microbial taxa present in the samples. These results confirm the high quality and reliability of the sequencing data, which are sufficient for downstream analyses.

3.2. ASV Analysis and Species Annotation

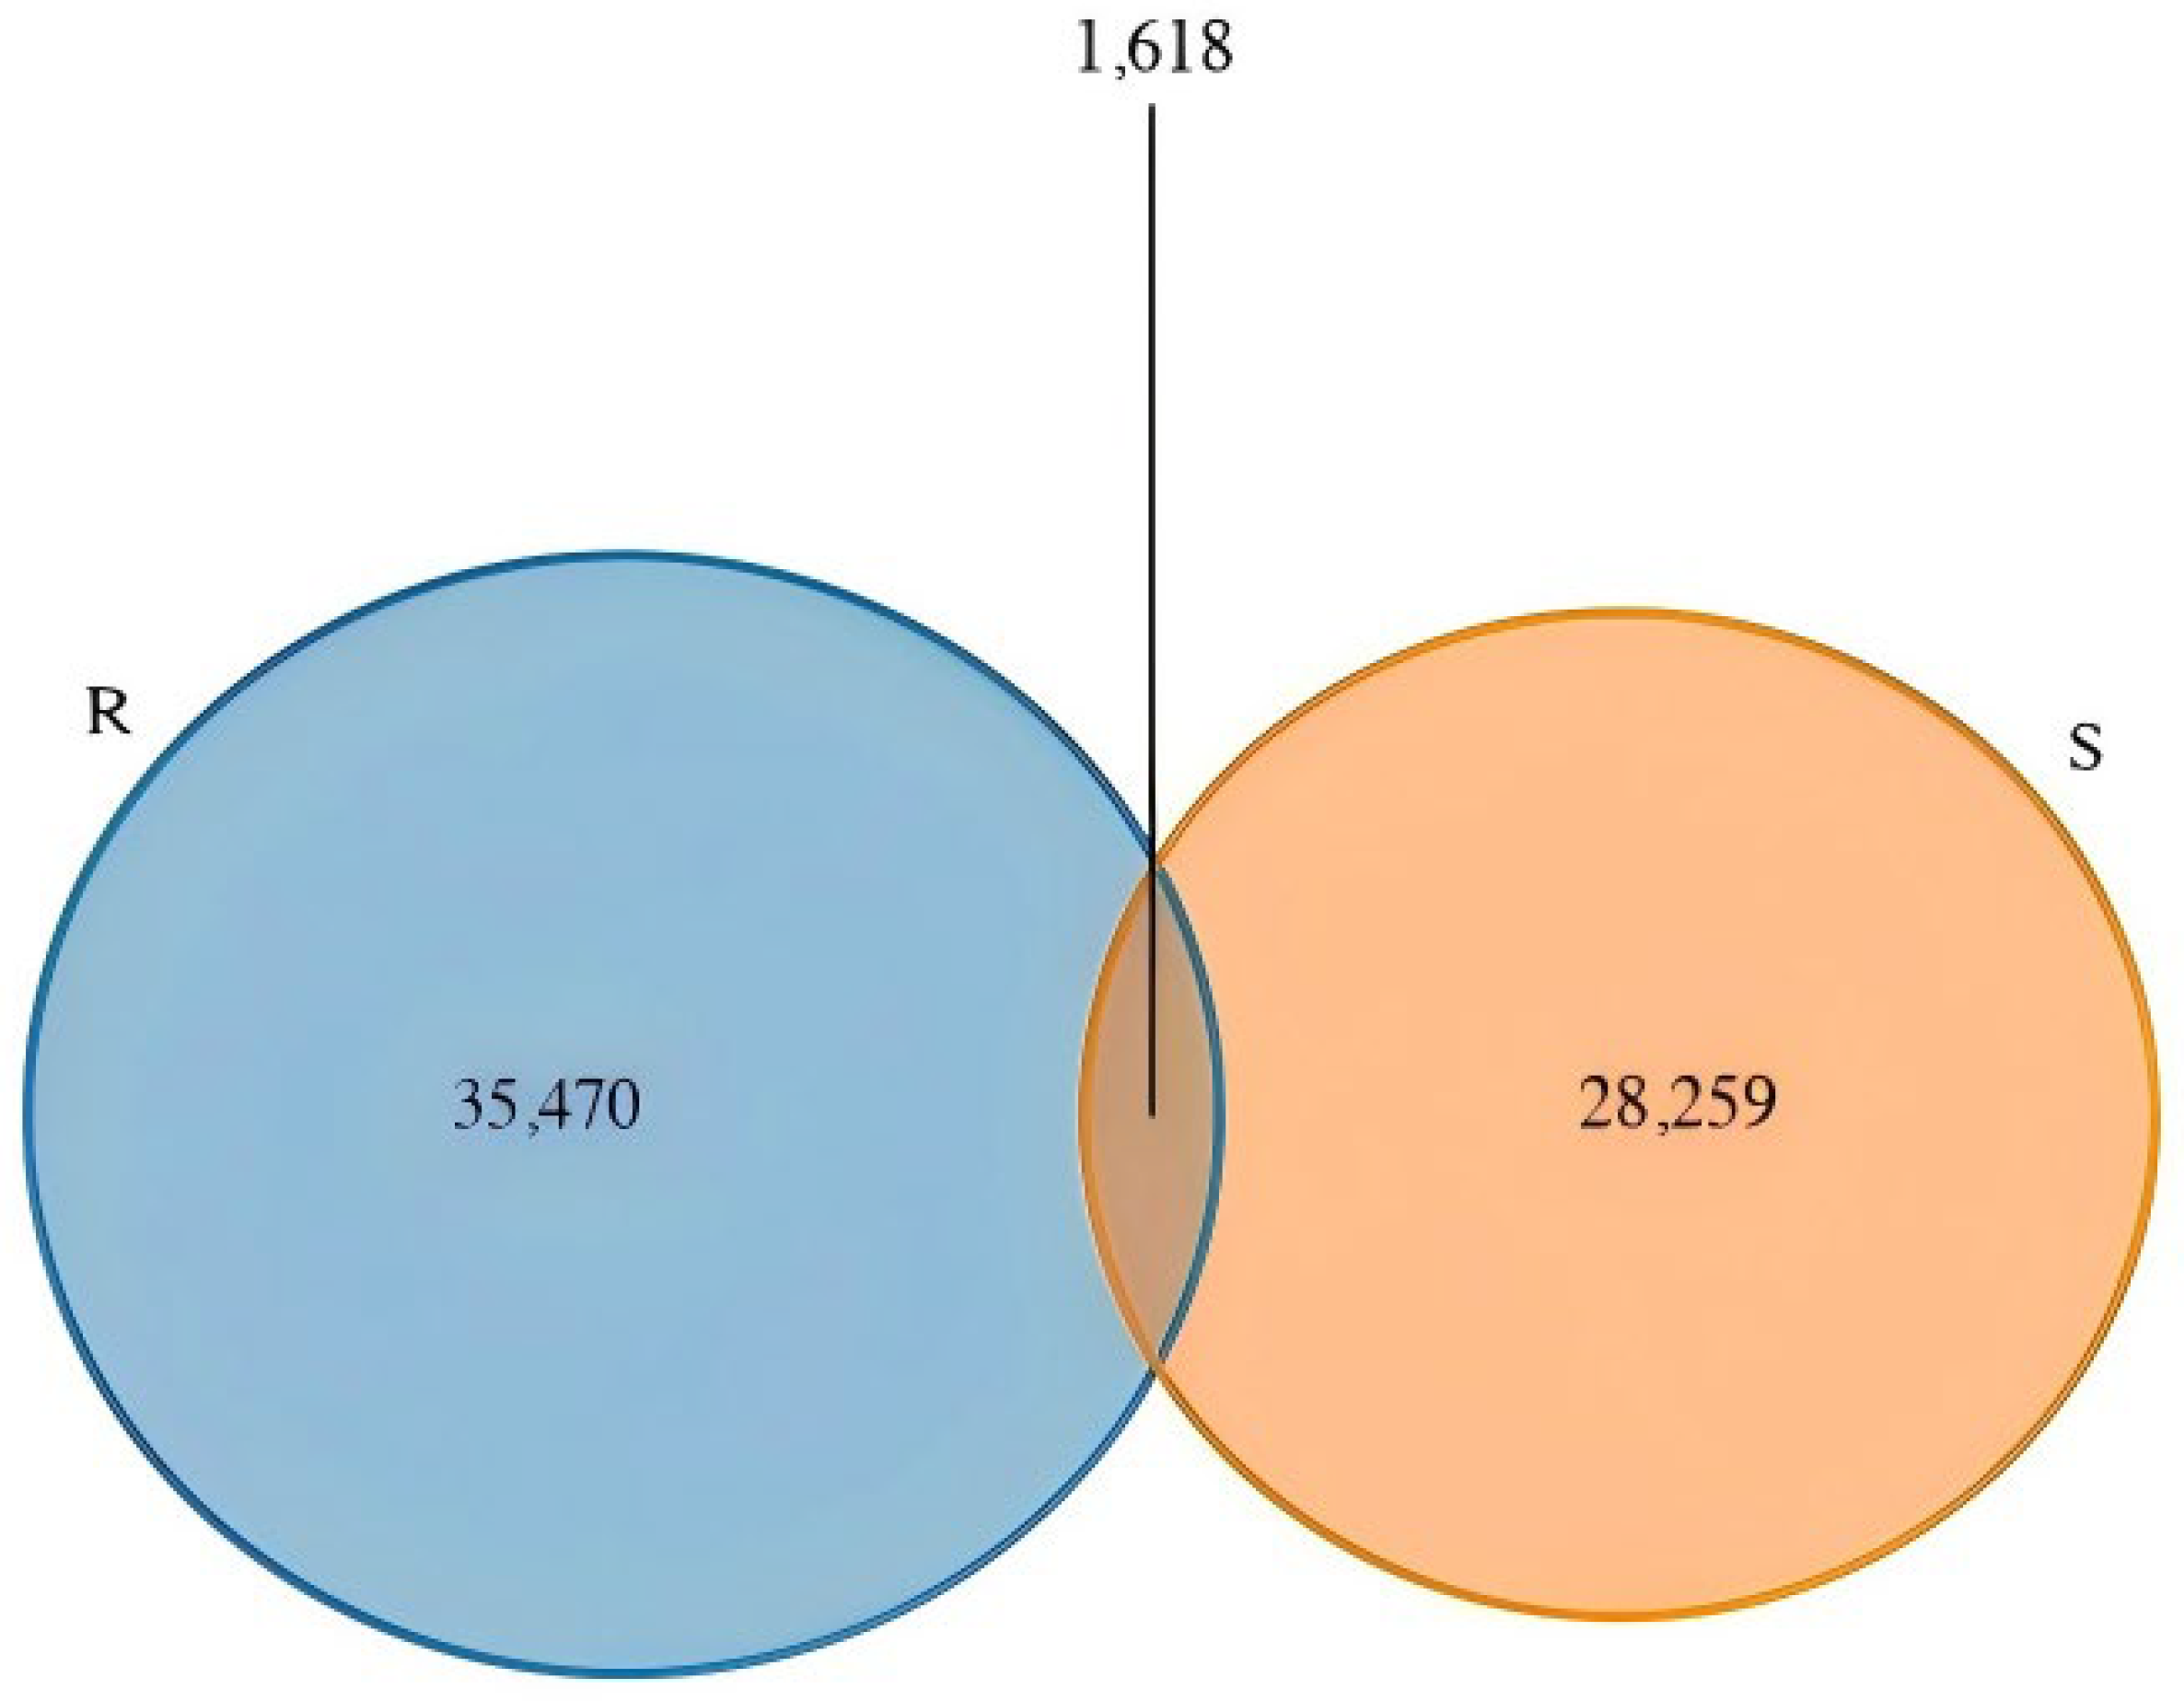
Based on DADA2-based denoising, a total of 65,347 amplicon sequence variants (ASVs) were identified. Based on the OTU (Operational Taxonomic Unit) analysis of the 30 soil samples, distinct microbial community profiles were observed between the two groups. Analysis of ASV distribution between the two sample groups revealed a core set of 1618 ASVs shared between the petroleum-contaminated (Group S) and bioremediated (Group R) sites, representing microbial taxa that persisted across both environmental conditions (Figure 3). The bioremediated site exhibited a substantially larger unique ASV repertoire of 35,470 ASVs, reflecting the high microbial diversity and ecological complexity recovered after phytoremediation. In contrast, the contaminated site harbored 28,259 unique ASVs, indicating the presence of specialized microbial taxa adapted to petroleum hydrocarbon stress. This pattern of ASV sharing and uniqueness demonstrates that while both environments maintain distinct microbial assemblages, phytoremediation has facilitated the establishment of a more diverse and functionally redundant microbial community in Group R.

3.3. Analysis of Microbial Community Composition

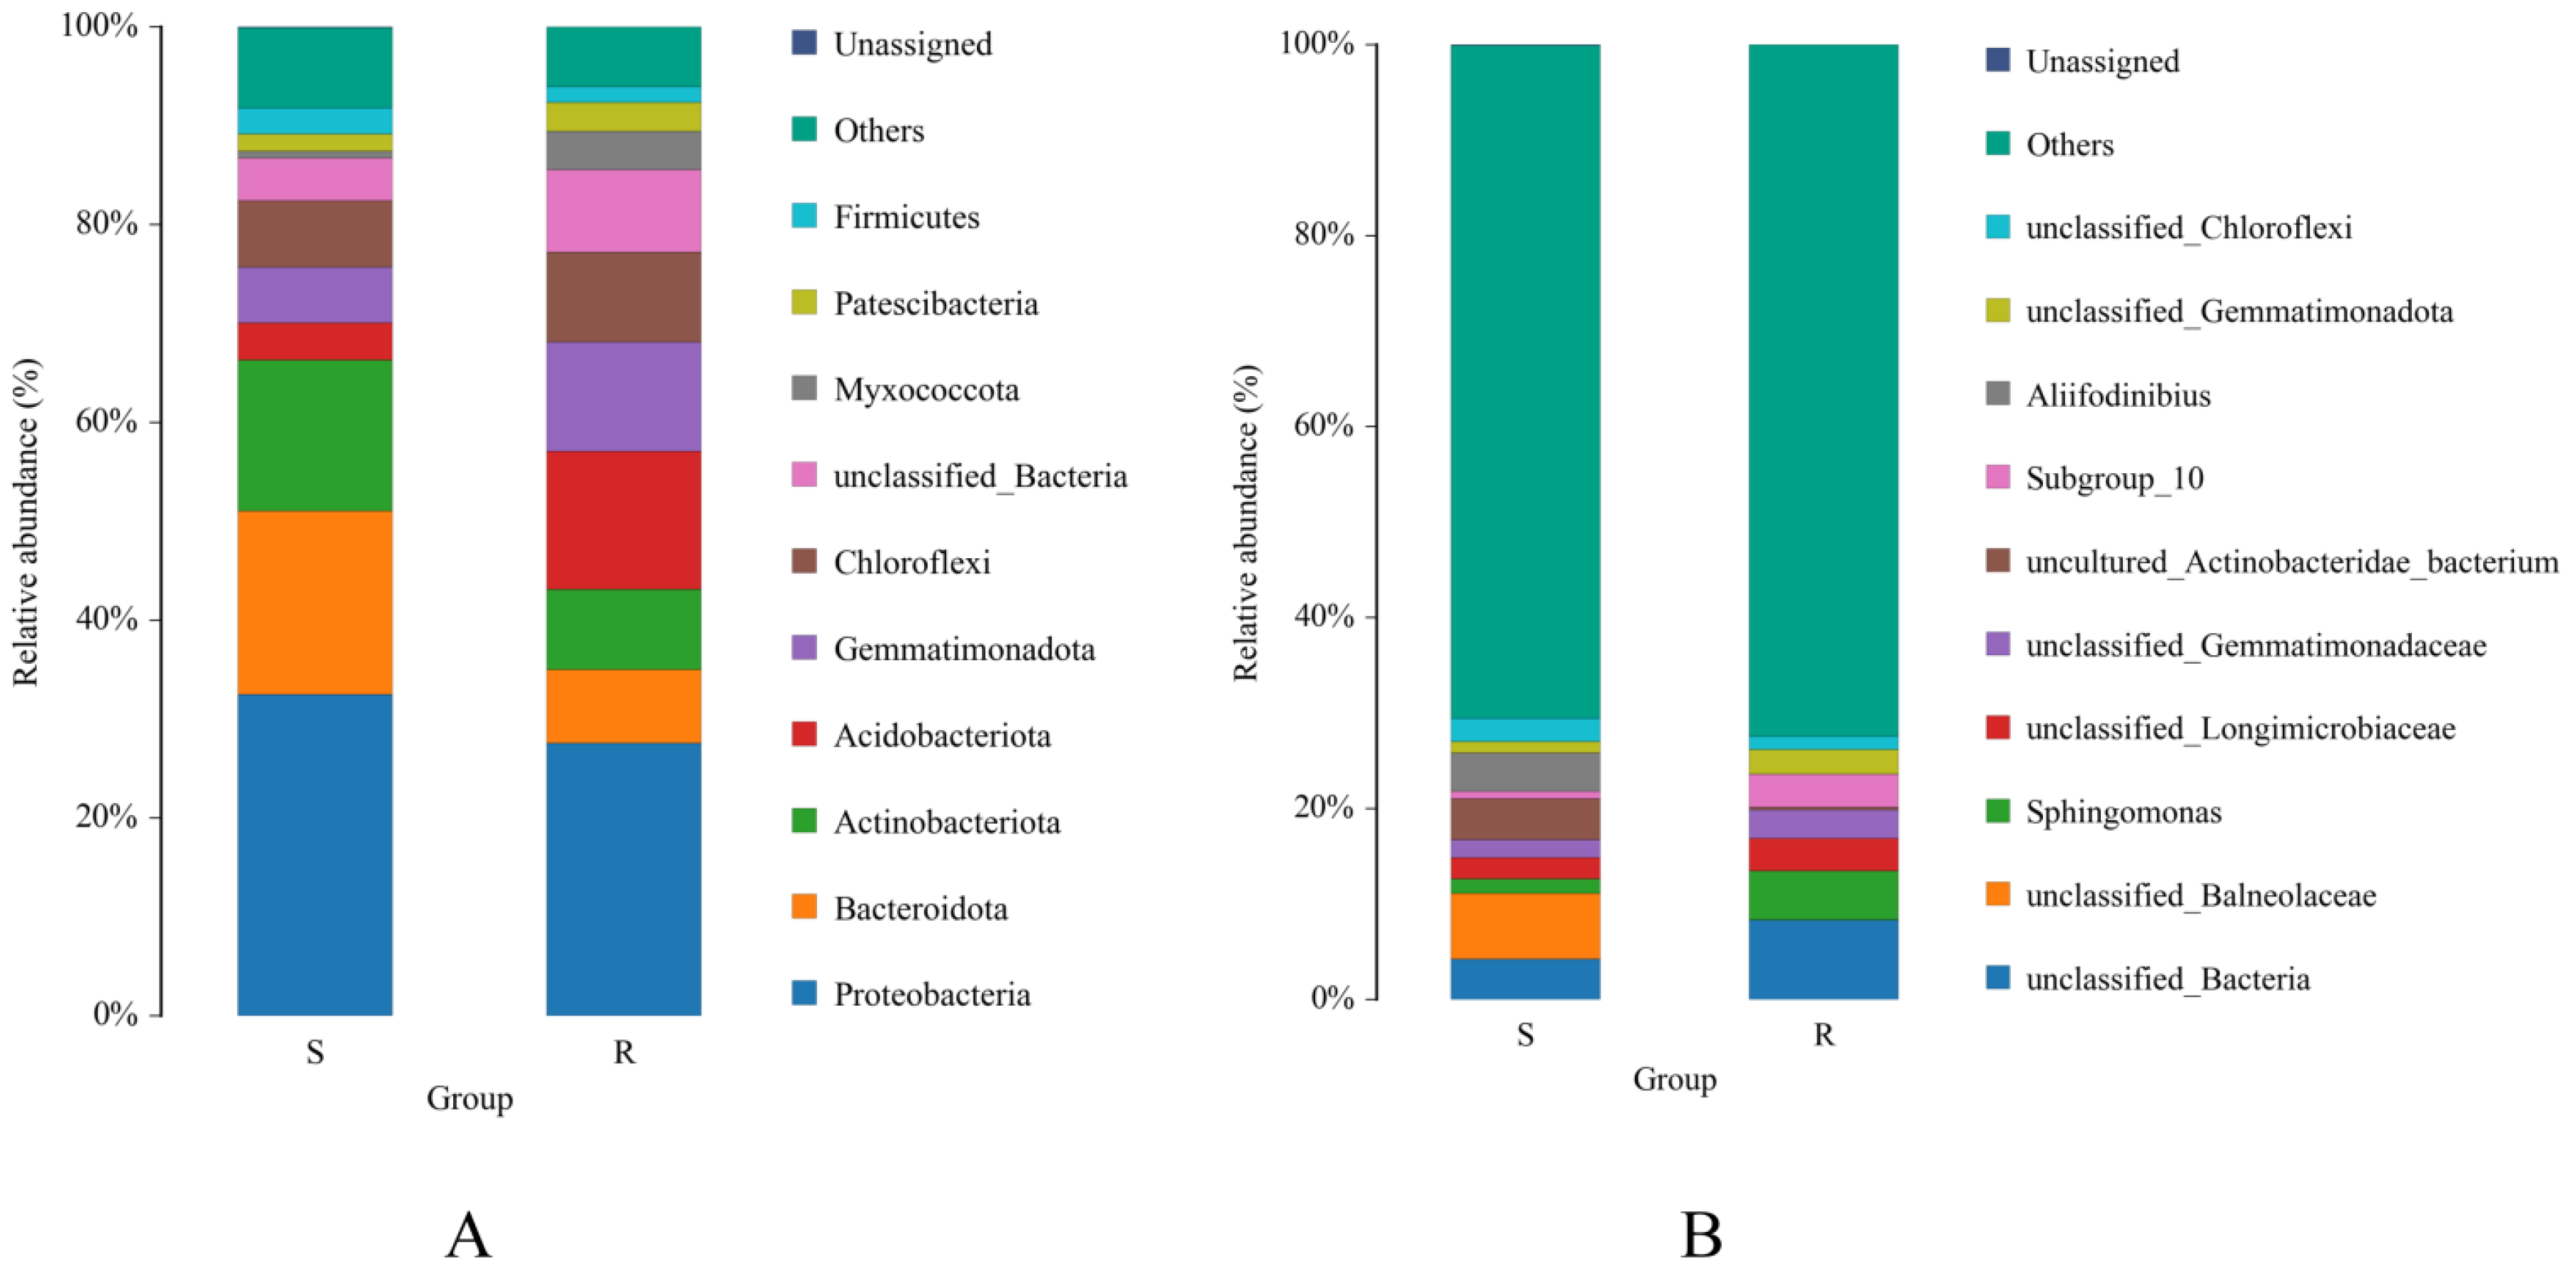
Based on the comparative analysis of the microbial community structures, a distinct inter-group divergence is clearly observed between the bioremediated (Group R) and petroleum-contaminated (Group S) soils. Group R is mainly characterized by a high relative abundance of hydrocarbon-degrading phyla, predominantly Pseudomonadota, while Bacteroidota and Actinobacteriota play supporting roles. This reflects a selectively enriched community that is well-adapted for pollutant breakdown. In contrast, Group S shows a more diversified phylogenetic profile, with a significantly higher prevalence of oligotrophic and stress-tolerant phyla such as Chloroflexi, Acidobacteria, and Firmicutes. This distinct shift in dominant phyla, from a specialized, catabolically active consortium in Group R to a survival-oriented, generalist community in Group S, effectively demonstrates the profound impact of contamination and bioremediation on soil microbial assembly.
Based on the genus-level taxonomic composition, significant inter-group differences are detected between the two sample types (Figure S1). The microbial community in group R is marked by a high relative abundance of Sphingomonas, a genus well-recognized for its function in polycyclic aromatic hydrocarbon degradation, along with considerable proportions of unclassified lineages from Longimicrobiaceae and Gemmatimonadaceae (Figure 4B). This indicates the existence of specialized but poorly characterized bacterial clades. In contrast, group S exhibits a distinct dominance of Alifodinibius, a halotolerant genus, along with a greater abundance of unclassified members of Chloroflexi and Gemmatimonadota, which typically thrive in nutrient-limited or stressed environments.

3.4. Alpha Diversity Analysis

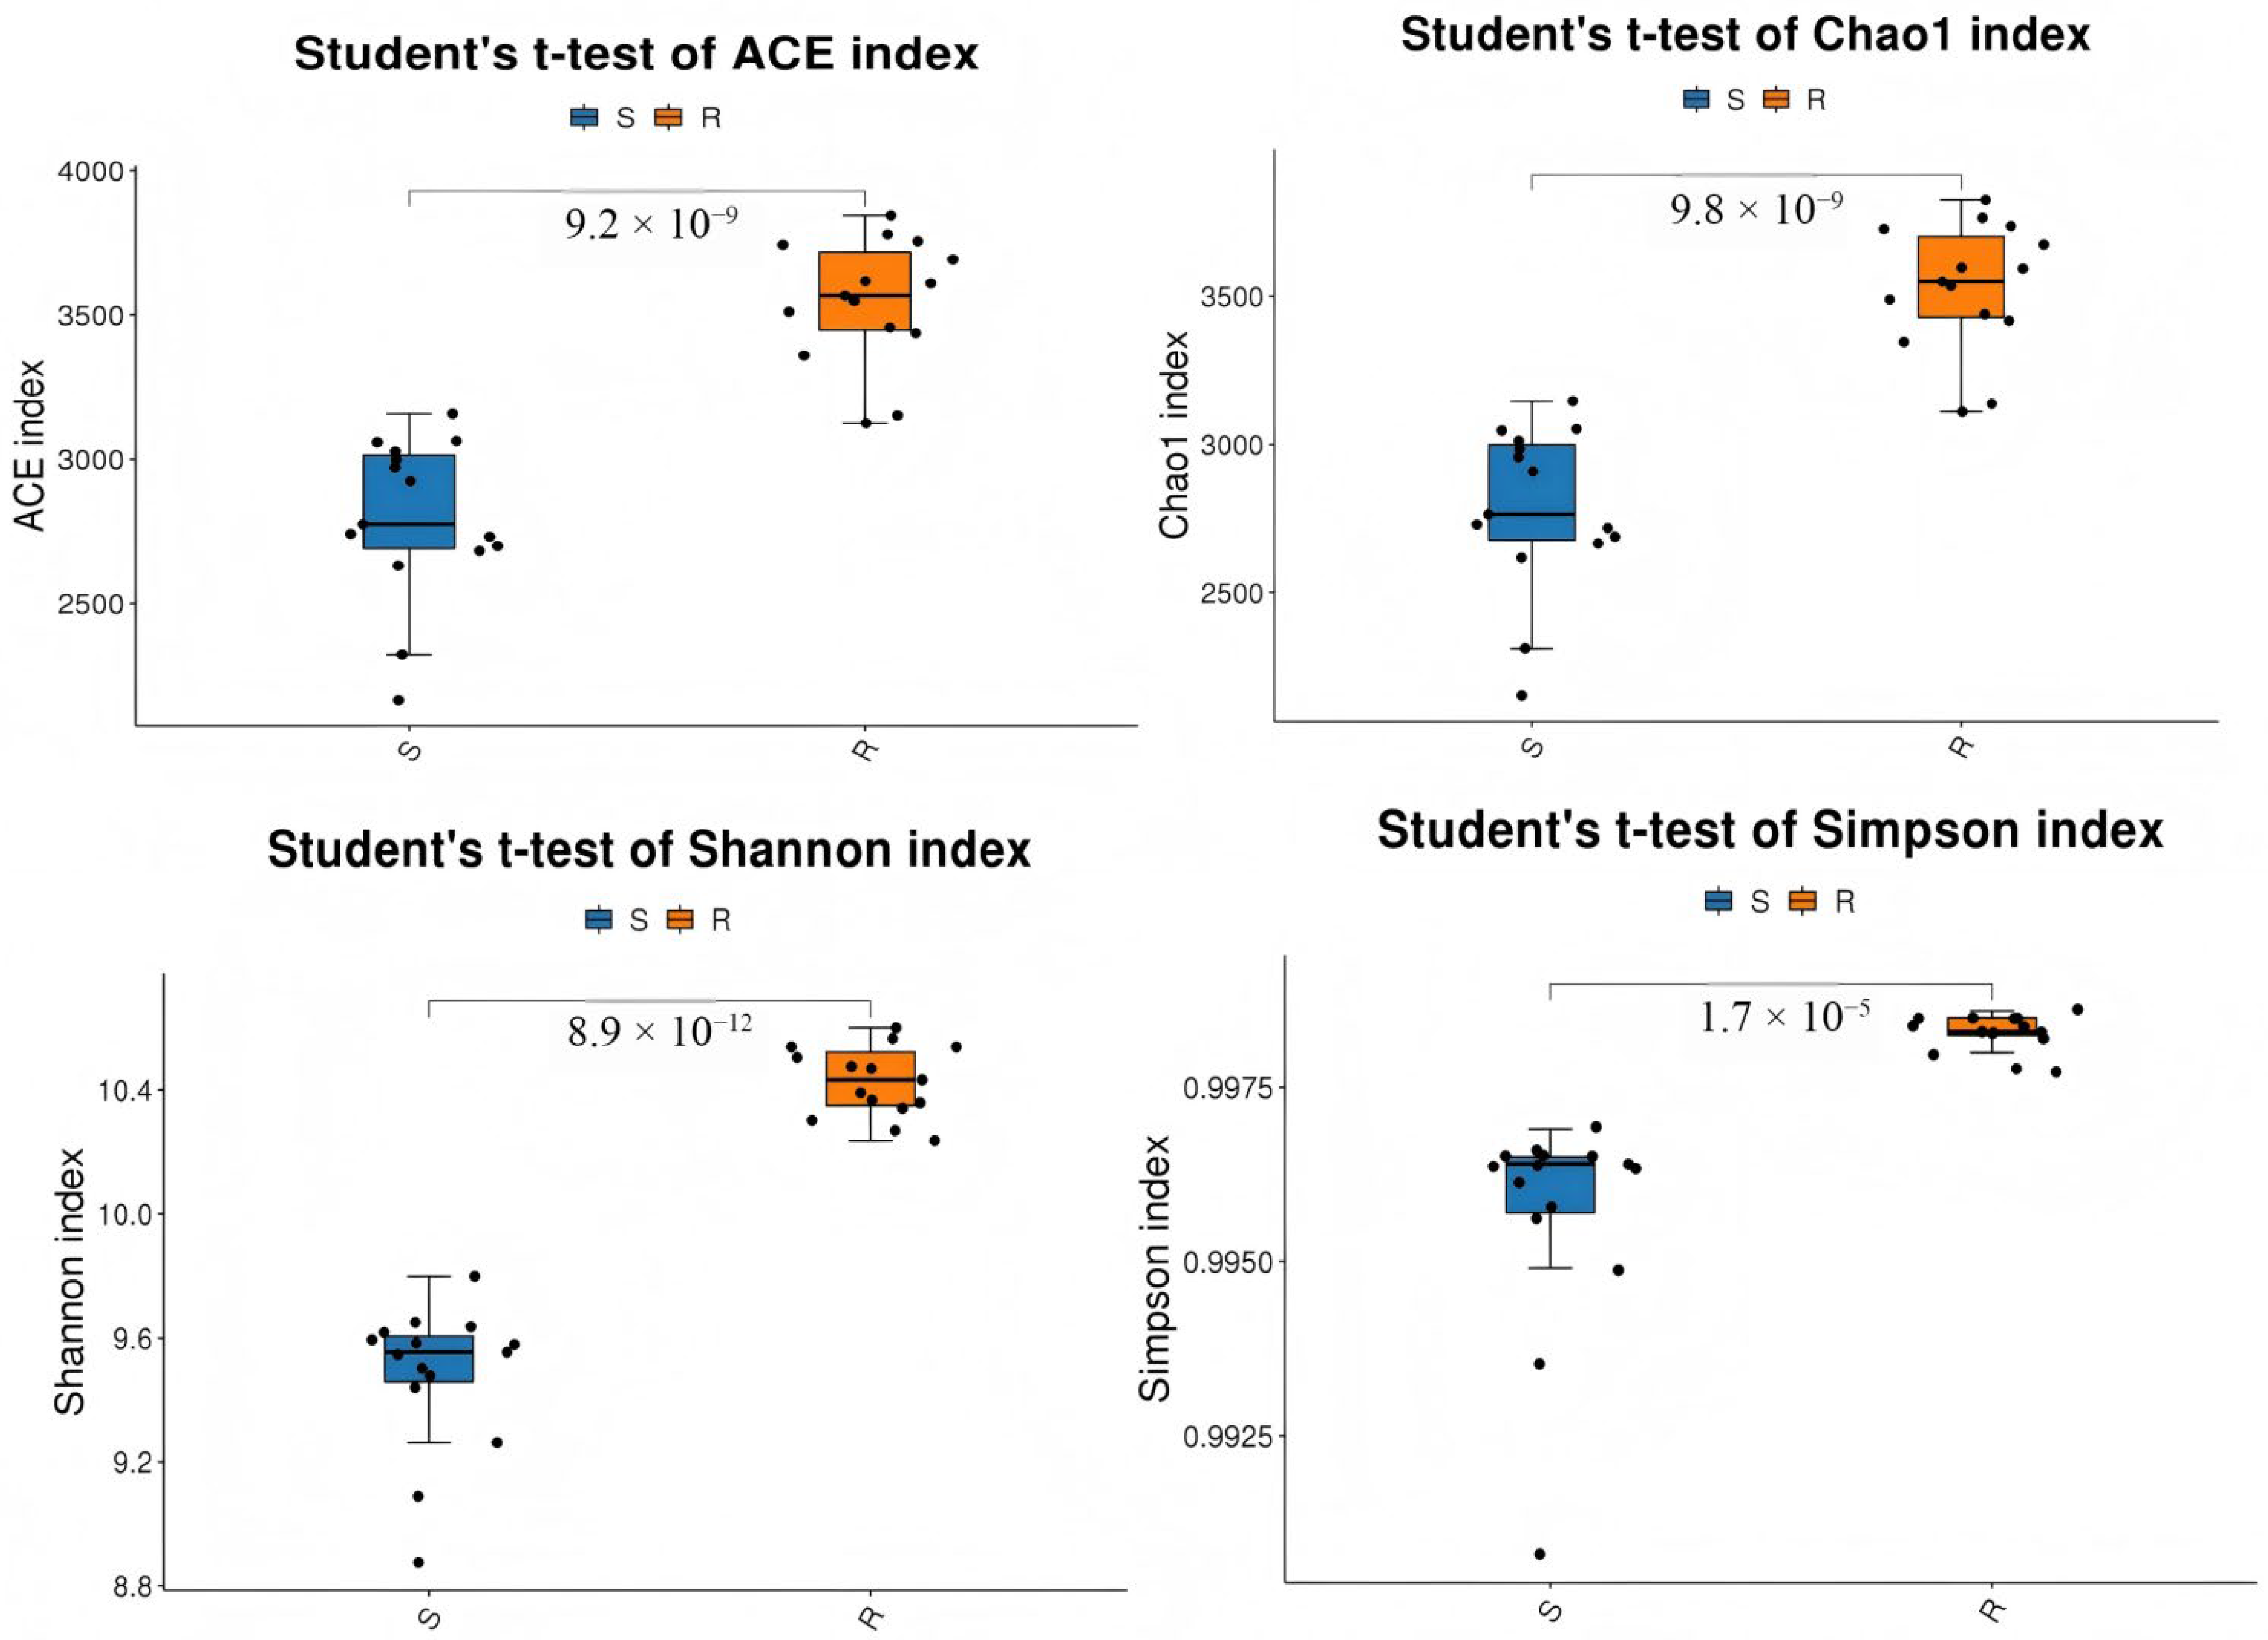
The highly significant differences in alpha diversity indices (p < 0.001) between the contaminated (S) and bioremediated (R) groups reveal a profound ecological transformation (Figure 5). The substantially higher richness (ACE, Chao1) and diversity (Shannon, Simpson) in Group R demonstrate that bioremediation has successfully fostered a more complex and robust microbial ecosystem. This shift from the impoverished community in contaminated soils to a rich, diverse consortium in remediated soils indicates not just quantitative recovery but qualitative ecological restoration.

3.5. Beta Diversity Analysis and Significance of Group Differences

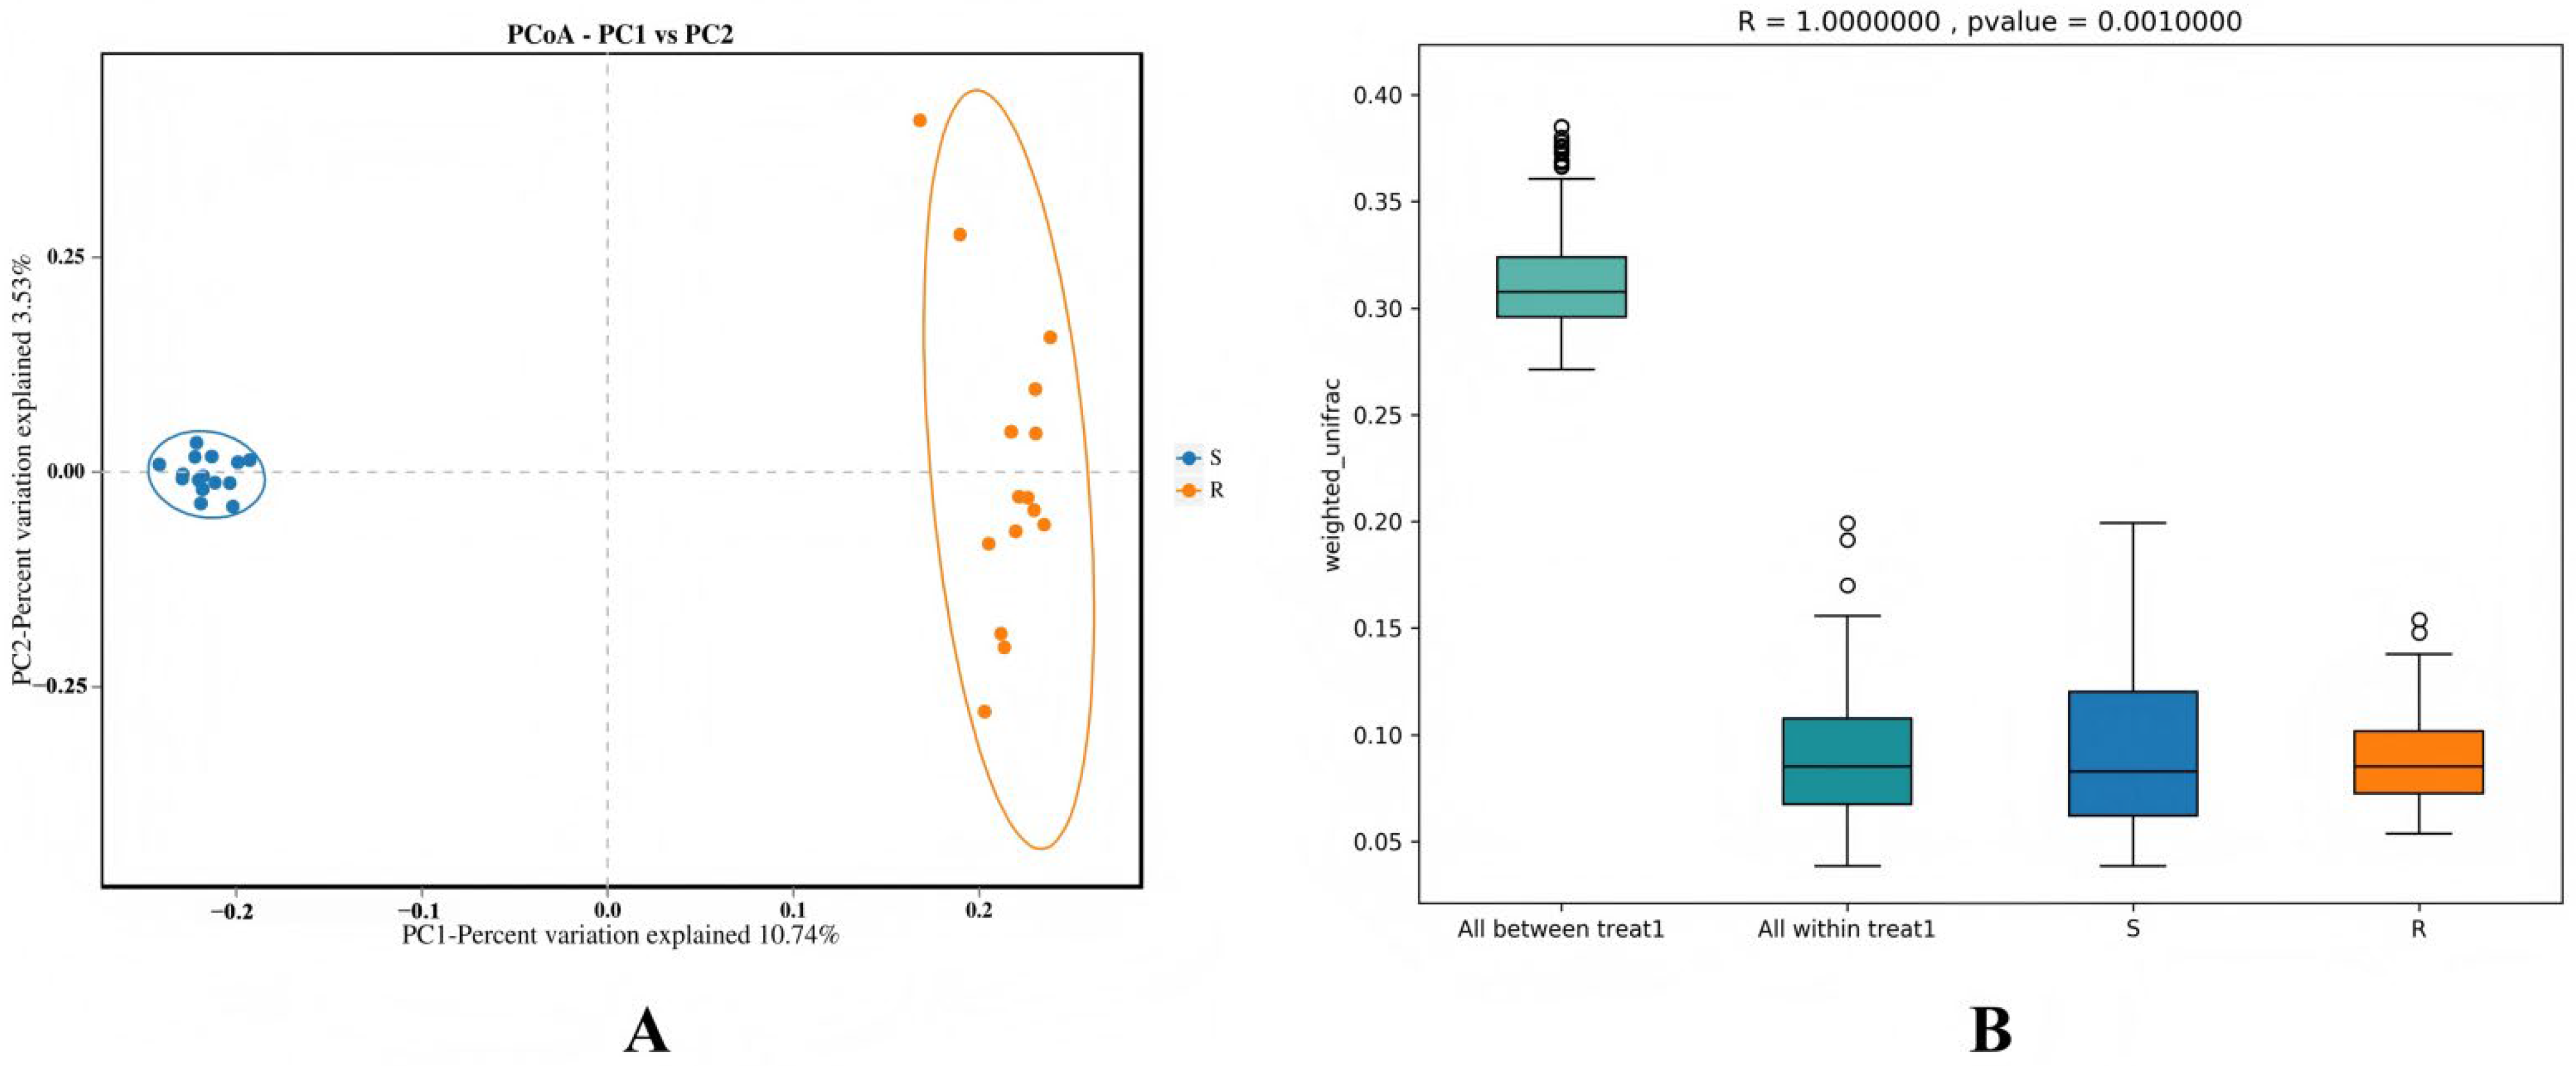
Based on the provided PCoA (Principal Coordinates Analysis) scatter plot, the data points are visualized along the first two principal components, PCoA1 and PCoA2 (Figure 6A). PCoA1 is the dominant component, explaining a substantial 10.74% of the total variation within the dataset. In contrast, PC2 accounts for a much smaller portion, explaining only 3.53% of the variation. The data points are distributed across a range of approximately −0.25 to 0.25 on the PCoA1 axis and from −0.2 to approximately −0.2 on the PCoA2 axis. This indicates that the primary structure and major patterns of the data are effectively captured along the PCoA1 axis. Analysis of similarities (ANOSIM) further supported these findings (Figure 6B), showing that differences between groups were significantly greater than those within groups (R = 1.0, p = 0.001). An R value of 1.0 represents the theoretical maximum of the ANOSIM statistic, indicating that all ranks within groups are smaller than any rank between groups. The UPGMA clustering dendrogram and sample clustering heatmap consistently grouped samples into two distinct clusters corresponding to the S and R groups.

3.6. Inter-Group Biomarker Analysis

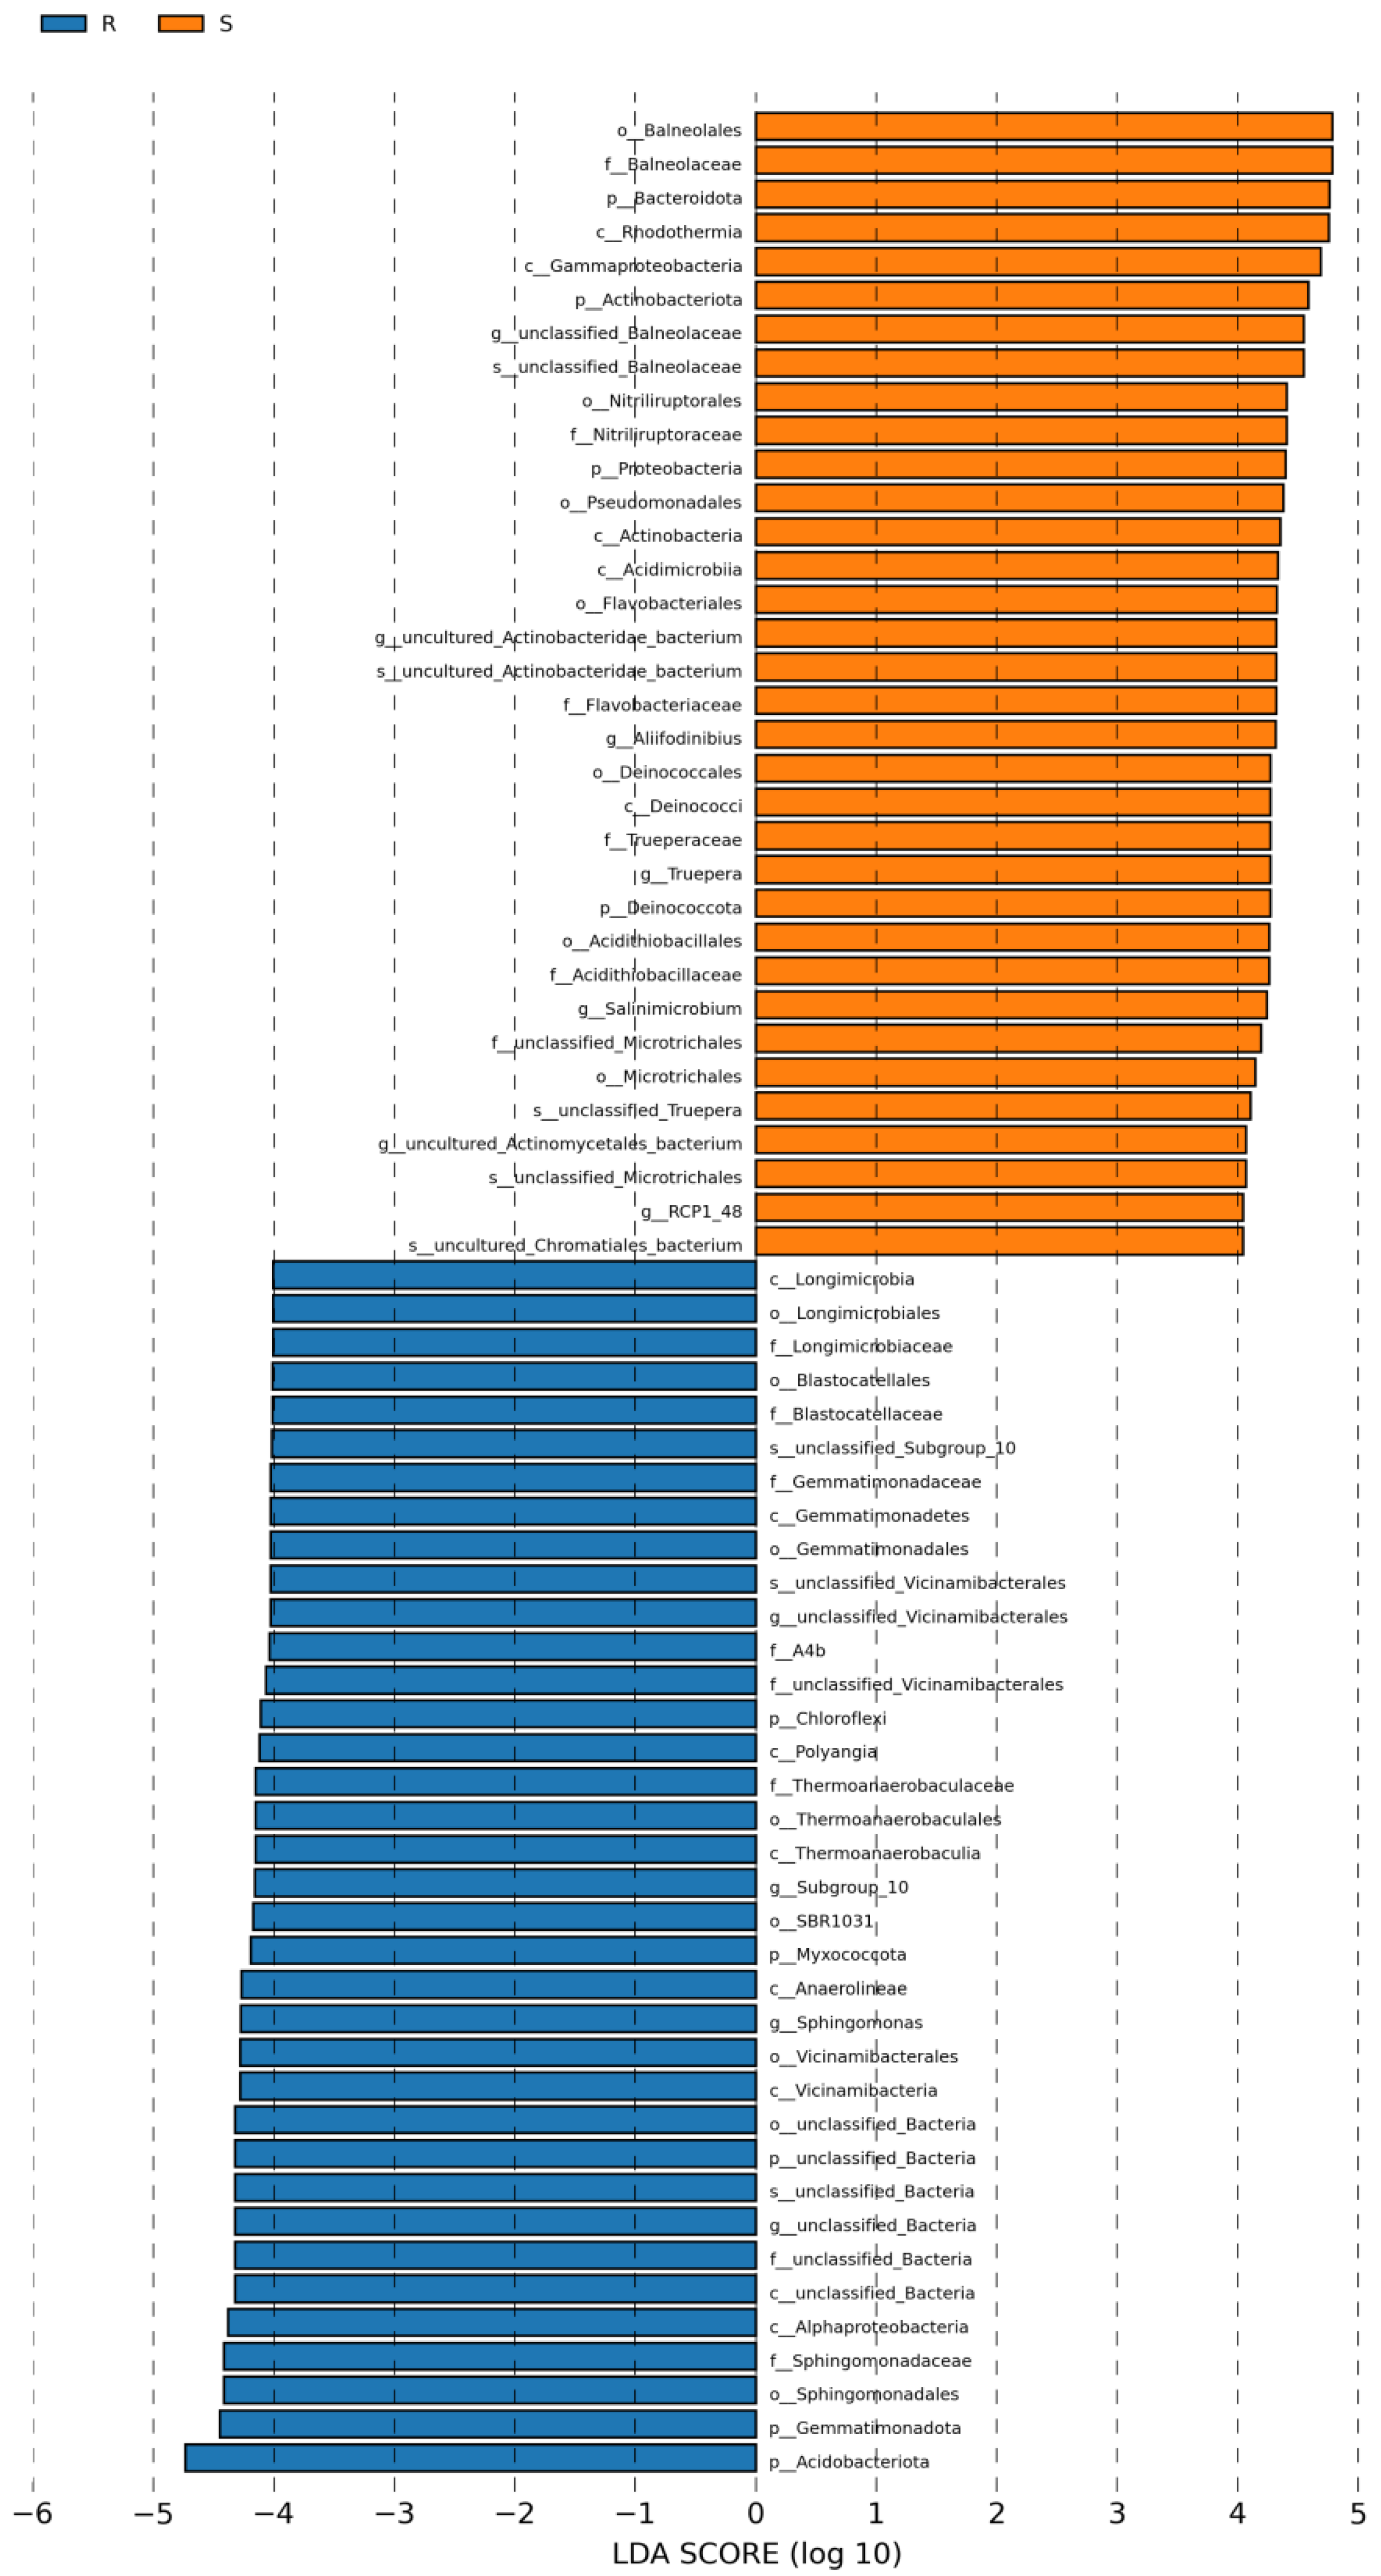
Based on the LEfSe analysis presented in the figure, distinct taxonomic biomarkers were identified for the petroleum-contaminated (Group S) and bioremediated (Group R) sites (LDA score > 4.0, p < 0.05). The cladogram and bar plot reveal that Group S (contaminated) is characterized by the enrichment of several well-known hydrocarbon-degrading and stress-tolerant taxa (Figure 7). Key biomarkers enriched in Group S include the phyla Deinococcus and unclassified members of Actinomycetes, as well as genera such as Truepera (phylum Deinococcota), Alifodinibius (phylum Balneolaceae), and Treponera. The enrichment of Gammaproteobacteria, Pseudomonadales, and Acidithiobacillales in Group S further supports the selective pressure of petroleum hydrocarbons, as these groups are frequently associated with xenobiotic degradation and adaptation to contaminated environments. In contrast, Group R (remediated) exhibits a distinct set of enriched taxa, including the phyla Pseudomonadota, Bacteroidota, and Actinobacteriota, along with specific orders such as Flavobacteriales, Nitrilirhopanales, and Microtrichales.
At the genus level, Sphingomonas is a genus well-documented for polycyclic aromatic hydrocarbon (PAH) degradation was significantly enriched in Group R, together with unclassified members of Balneolaceae and Actinomycetes. This taxonomic shift reflects the recovery of a more diverse and metabolically versatile microbial community following phytoremediation, consistent with the functional predictions indicating enhanced nutrient cycling and reduced pollution stress in remediated soils. Overall, the LEfSe analysis confirms that petroleum contamination selects for a specialized microbiome enriched in hydrocarbon degraders and stress-tolerant taxa, while phytoremediation fosters the reassembly of a broader, functionally diverse microbial consortium indicative of ecological restoration.

3.7. Functional Prediction Analysis

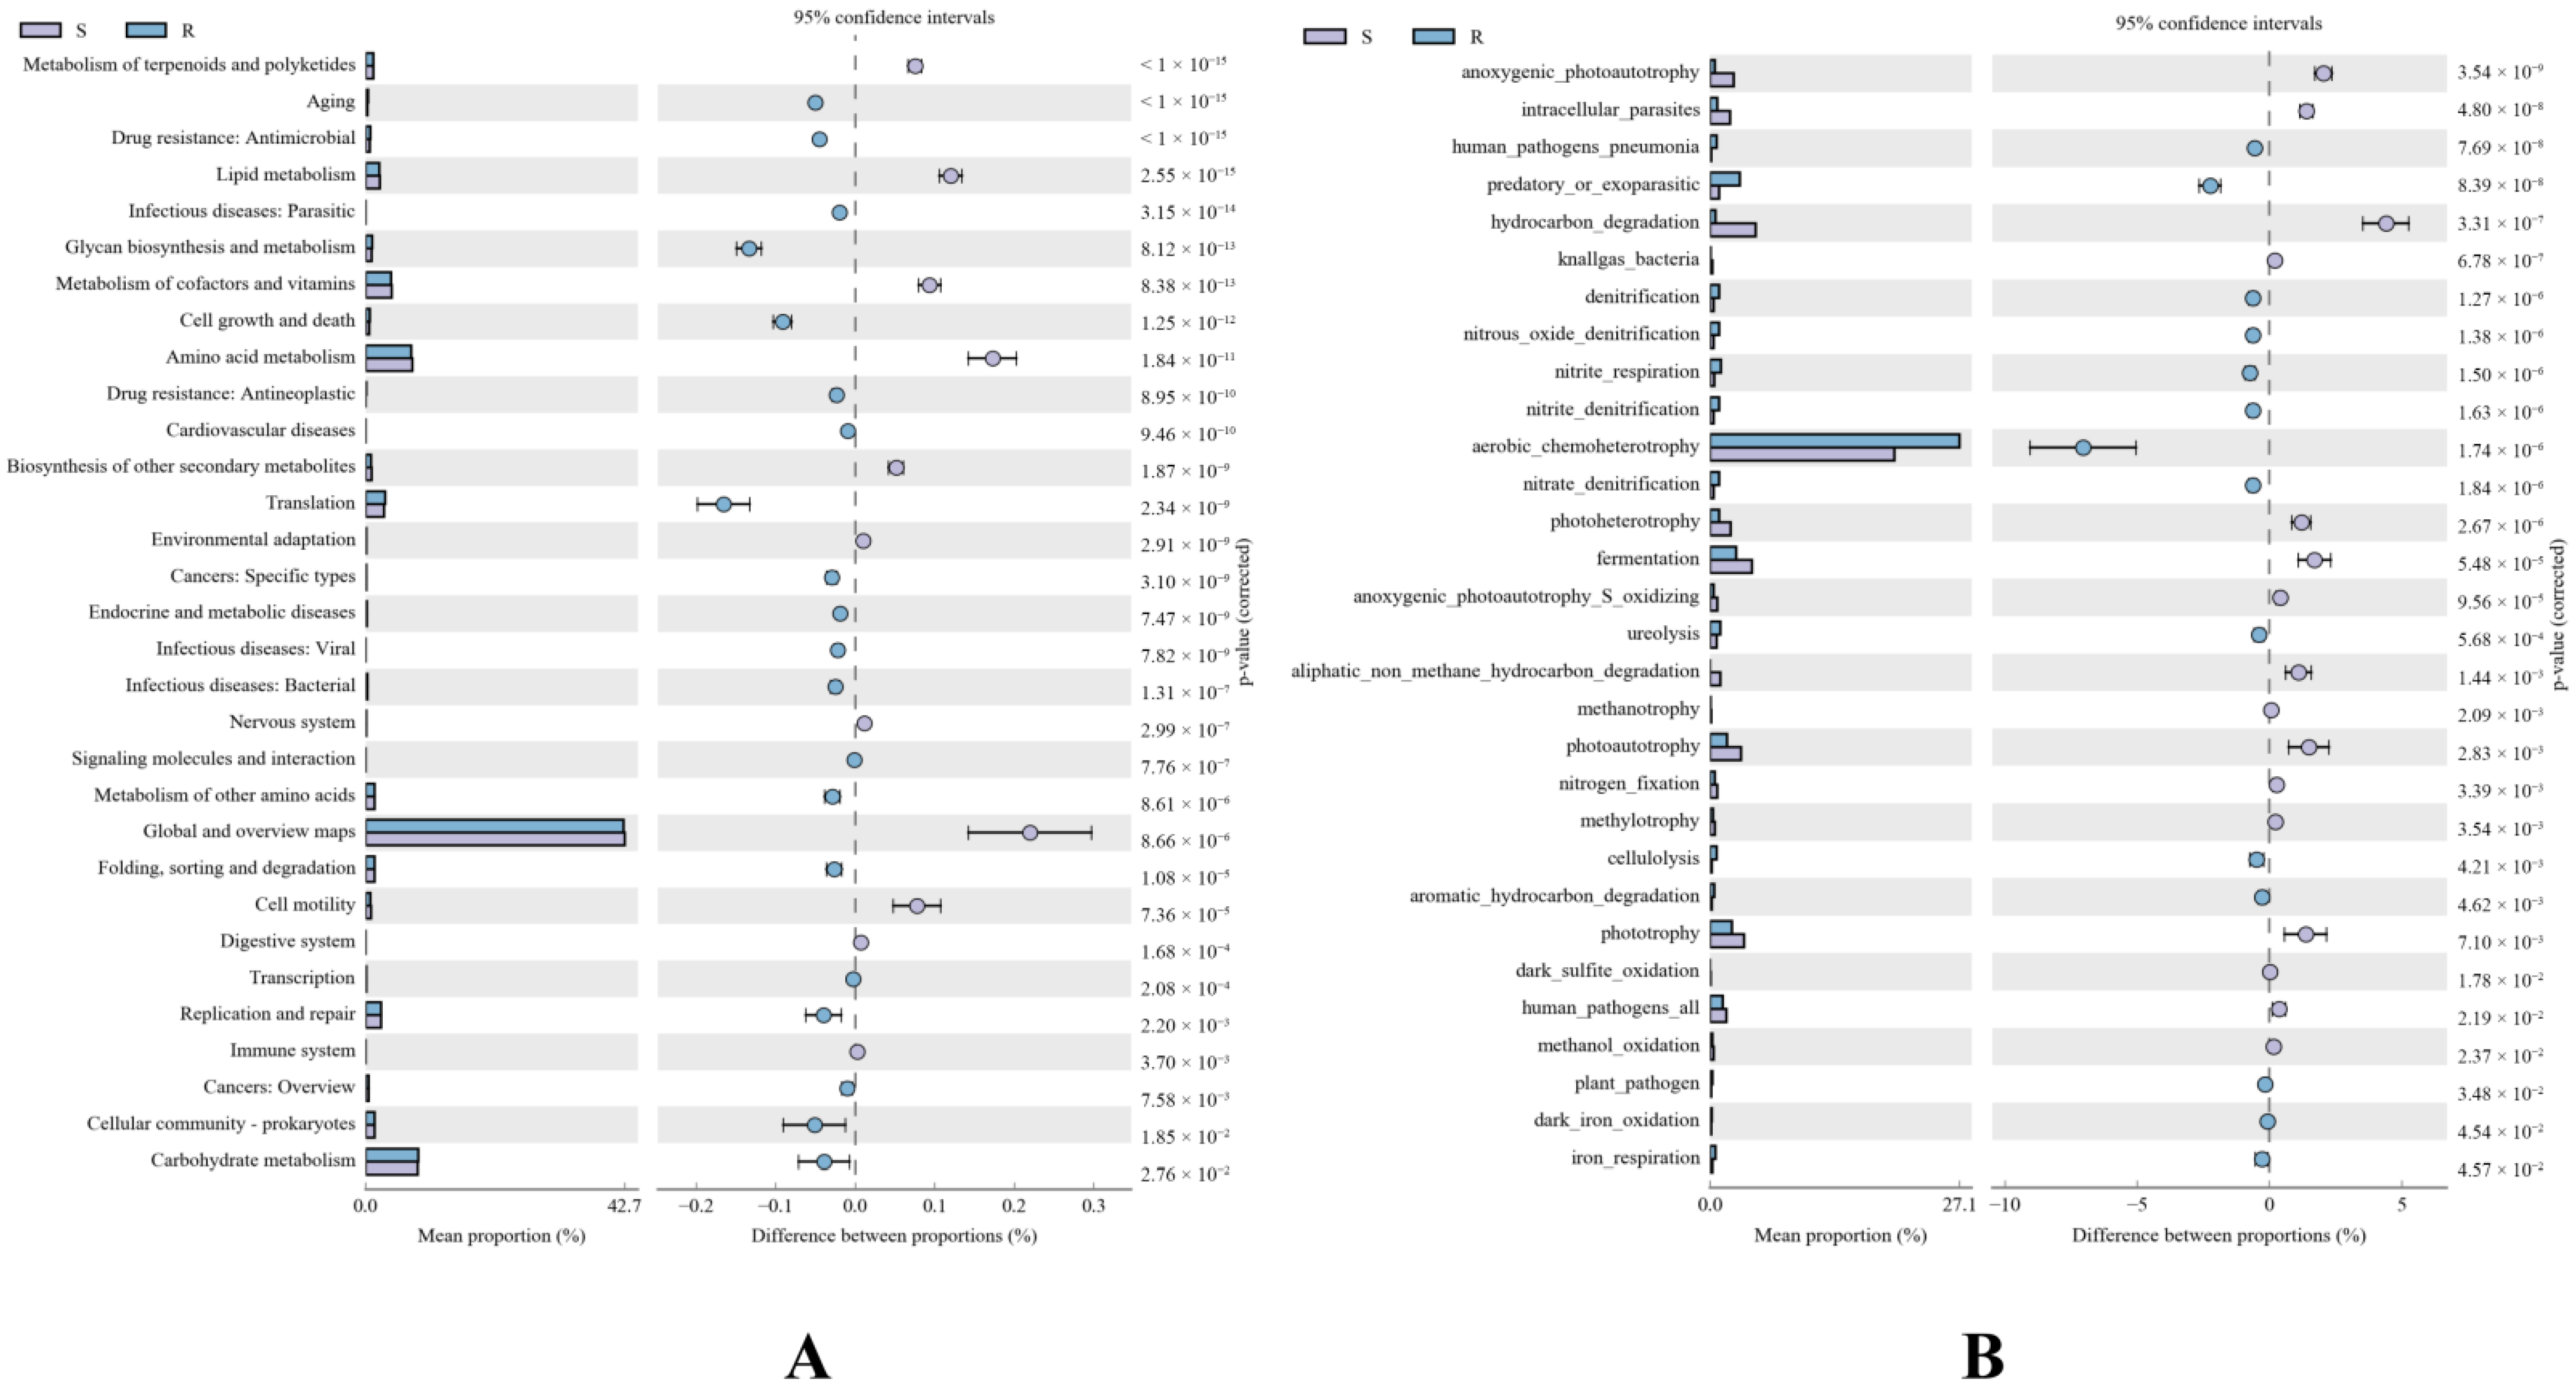
Functional profiles of microbial communities were inferred using PICRUSt2 v2.4.1 based on ASV phylogeny. Significant differences in predicted KEGG pathway abundances were observed between Group S and Group R (Figure 8A). Group S showed significant enrichment of pathways associated with “xenobiotics biodegradation and metabolism” (KEGG Level 1) and specific hydrocarbon degradation pathways. Functional predictions via PICRUSt2 reveal significant enrichment of KEGG pathways associated with “hydrocarbon degradation” (including aromatic and aliphatic hydrocarbon degradation) and “xenobiotic biodegradation and metabolism” in the S group (p-value < 0.05). Additionally, enhanced representation of pathways related to membrane transport and stress response suggests microbial adaptation to the contaminated environment. Conversely, the R group displays higher relative abundances in core metabolic pathways such as “amino acid metabolism,” “carbohydrate metabolism,” “energy metabolism,” and “signal transduction,” indicating a functional profile more representative of unpolluted soil ecosystems. It should be noted that these functions represent predicted functional potential based on community composition, rather than direct empirical evidence.
FAPROTAX-based analysis further clarifies the ecological roles of microbial communities in environmental settings (Figure 8B). The S group exhibits significantly higher predicted abundances for functions such as “petroleum hydrocarbon degradation,” “aromatic compound degradation,” “methylotrophy,” and “methanol oxidation,” suggesting a potential enhancement of hydrocarbon transformation capacity in the polluted soil microbial community (p-value < 0.05). In contrast, the R group shows an increased prevalence of processes associated with nitrogen and sulfur cycling, like “nitrification,” “denitrification,” and “sulfur respiration,” emphasizing the restoration of diverse biogeochemical functions in remediated areas.

4. Discussion

The structural and functional shifts observed in the microbial community, specifically the enrichment of hydrocarbon-degrading taxa in contaminated soils and the recovery of diverse metabolic pathways in remediated soils, suggest that microbial–plant collaborative remediation has the potential to enhance the removal of total petroleum hydrocarbons (TPH) and promote the restoration of soil ecological functions by improving the rhizosphere microenvironment and activating key functional genes. In saline–alkaline environments like the Yellow River Delta, Pseudomonadota, Actinomycetes, and Bacillus become the dominant microbial taxa in contaminated soils, while Acidobacteria show a reduced abundance in severely polluted areas [10,20]. Our results firmly corroborated that Pseudomonadota and Actinobacteriota were the predominant phyla across both contaminated and remediated soils. This finding implies that these phyla might encompass stress-tolerant lineages capable of persisting under petroleum hydrocarbon contamination in the Yellow River Delta. At the genus level, Bacillus, a taxon commonly linked to hydrocarbon degradation, displayed comparable abundances between the groups. This suggests its potential function as a generalist degrader in both environments. The present study systematically investigated the structural and functional responses of soil microbial communities to crude oil contamination and subsequent phytoremediation in the Yellow River Delta, a region of significant ecological and economic importance. Through high-throughput sequencing of the 16S rRNA gene and comprehensive bioinformatic analyses, we clearly demonstrated the substantial alterations in microbial diversity, community composition, and functional potential between contaminated (Group S) and remediated (Group R) soils. Our findings not only validate previous studies on microbial adaptation to hydrocarbon stress but also provide novel perspectives on the restoration dynamics of soil ecosystems after bioremediation.
Our results demonstrate that petroleum contamination exerts a strong selective pressure on soil microbial communities, leading to a significant reduction in alpha diversity. This decline is consistent with the well-documented toxicity of petroleum hydrocarbons, which inhibit the growth of sensitive microorganisms while favoring the proliferation of tolerant or hydrocarbonoclastic taxa [21,22]. The observed simplification of community structure in Group S reflects environmental filtering, wherein only a subset of microorganisms capable of surviving in contaminated conditions persists. Beta diversity analyses, including PCoA, further confirmed the stark compositional divergence between Groups S and R, underscoring the role of petroleum pollutants as a dominant ecological filter that restructures microbial assemblages [13,23]. At the phylum level, contaminated soils were enriched with Chloroflexi, Acidobacteria, and Firmicutes-groups often associated with oligotrophic and stressed environments [24]. In contrast, remediated soils exhibited a higher relative abundance of Pseudomonadota, Bacteroidota, and Actinobacteriota, which are frequently linked to nutrient-rich conditions and active organic matter degradation. This shift suggests that contamination favors stress-tolerant specialists, while remediation encourages the recovery of metabolically versatile taxa. LEfSe analysis revealed that genera such as Alcanivorax, Marinobacter, and Truepera were significantly enriched in contaminated soils (Group S) (Figure 7). These taxa are well-documented hydrocarbon degraders [25,26], and their co-occurrence suggests the formation of a functional consortium adapted to petroleum contamination. The presence of unclassified lineages within Gammaproteobacteria and Bacteroidota also points to potentially novel degraders that warrant further investigation [27]. These findings highlight the need for further taxonomic and functional characterization of these cryptic microbial groups.
The remediation process resulted in a substantial recovery of microbial diversity and complexity within Group R. The elevated values of the Chao1, ACE, Shannon, and Simpson indices in Group R demonstrate that phytoremediation not only diminished pollutant levels but also reinstated ecological conditions favorable for microbial recolonization and growth. The augmentation of microbial richness and evenness implies a transformation from a specialized, stress-adapted community to a more diverse and functionally redundant one, which is typically linked to enhanced ecosystem stability and resilience. Functional predictions carried out using PICRUSt2 disclosed an evident shift in metabolic potential between the two groups. In Group S, pathways associated with hydrocarbon degradation, xenobiotic metabolism, and stress response were dominant, mirroring a community adapted to survival in a contaminated environment. In contrast, the phytoremediated soils (Group R) displayed significantly greater alpha diversity and a more uniform community structure, indicating a transition from a stress-adapted to a metabolically versatile microbial assemblage. Functional predictions additionally supported this shift, as Group R showed an enrichment in pathways related to core metabolism [12,28], while Group S was dominated by xenobiotic degradation and stress-related functions. These alterations suggest a restoration of ecosystem multifunctionality subsequent to remediation.
The microbial community patterns observed in this study are consistent with global trends in petroleum-contaminated soils, yet they also reflect the unique environmental context of the Yellow River Delta. The region’s saline–alkaline soils, seasonal hydrology, and history of oil extraction likely interact to shape the assembly and function of hydrocarbon-degrading microbiota. For instance, the enrichment of halotolerant genera such as Alifodinibius in Group S (Figure S1) may be attributed to the combined stress of salinity and hydrocarbon exposure, a feature less common in non-saline contaminated sites [13]. Similarly, another study found strong correlations between soil physicochemical parameters and microbial community structure, reporting considerable spatial variation in both alpha diversity and taxonomic turnover across contaminated sites under different environmental gradients [29]. Collectively, these findings imply that the characteristic sedimentary depositional environment, saline–alkaline soil matrix, and dynamic hydrological regime of the Yellow River Delta jointly restrict and regulate the assembly of hydrocarbon-degrading microbiota, thus explaining the region-specific patterns observed in this study. The recovery of microbial diversity in Group R may be partly attributed to rhizosphere effects mediated by Cynodon dactylon, the dominant plant at the remediated site. Root exudates likely provided labile carbon and energy sources, alleviating nutrient limitations and stimulating microbial activity [12]. This interpretation is consistent with the increased abundance of copiotrophic phyla (e.g., Pseudomonadota, Bacteroidota) in Group R and the enrichment of Sphingomonas, a genus known for PAH degradation and plant association.
The remarkable successful recovery of microbial diversity and function in Group R strongly underscores the effectiveness of phytoremediation as a sustainable strategy for restoring oil-contaminated soils. The plant-microbe interactions facilitated by phytoremediation presumably improved soil structure, nutrient availability, and microclimate, thus effectively supporting microbial reassembly and functional recovery [12,13]. These findings hold significant implications for the design and monitoring of bioremediation projects in the Yellow River Delta and other oil-impacted regions. Moreover, the identification of key hydrocarbon-degrading taxa and their associated functional traits offers a solid basis for developing targeted bioaugmentation strategies. The identification of key hydrocarbon-degrading taxa such as Alcanivorax and Marinobacter in contaminated soils offers a basis for developing targeted bioaugmentation strategies. These organisms could be explored as candidates for enhancing initial oil degradation, while phytoremediation may support long-term ecological restoration. A limitation of this study is the lack of accompanying soil physicochemical data (e.g., TPH concentration, pH, salinity), which would have enabled direct correlation with microbial community shifts. Future studies should integrate geochemical measurements with high-throughput sequencing to establish causal links and validate the ecological recovery processes inferred here.

5. Conclusions

This study conducted a systematic comparison of the differences in soil microbial communities between oil-contaminated and remediated areas in the Yellow River Delta. The key findings are as follows: Oil pollution led to a significant reduction in soil microbial alpha diversity and a profound alteration of the community structure. This resulted in a distinct separation between the microbial communities of the contaminated and remediated sites, which was verified by beta-diversity analyses (PERMANOVA, ANOSIM) and ordination methods. In contaminated soils, specific hydrocarbon-degrading bacterial taxa were enriched, including genera such as Alcanivorax and Marinobacter. These taxa act as crucial biomarkers in response to petroleum contamination. They were identified through LEfSe analysis and validated by Metastats, offering strong evidence for the microbial adaptation to pollution stress. The observed compositional and functional shifts, from a stress-adapted, hydrocarbon-degrading consortium in contaminated soils to a diverse, metabolically versatile community in remediated soils, clarify the profound impacts of petroleum pollution on soil ecosystems from the perspective of microbial ecology. Moreover, they reveal the dynamic changes in microbial communities during the pollution adaptation and remediation processes. These findings offer essential scientific evidence for assessing ecological risks in contaminated soils, screening efficient biodegradation agents, and guiding phytoremediation practices in the Yellow River Delta and similar saline–alkaline wetland ecosystems.

Supplementary Materials

The following supporting information can be downloaded at: https://www.mdpi.com/article/10.3390/d18040206/s1, Figure S1: The heatmap of the top 30 genera across all samples.

Author Contributions

H.W.: conceptualization, methodology, validation, formal analysis, investigation, resources, data curation, writing—original draft preparation, and writing—review and editing; S.L.: conceptualization, methodology, validation, formal analysis, investigation; X.T.: writing—original draft preparation, and writing—review and editing; L.Z.: conceptualization, methodology, validation, formal analysis, investigation, resources, data curation, writing—original draft preparation, and writing—review and editing. All authors have read and agreed to the published version of the manuscript.

Funding

This research received no external funding.

Institutional Review Board Statement

Not applicable.

Data Availability Statement

The original contributions presented in this study are included in the article/Supplementary Materials. Further inquiries can be directed to the corresponding author.

Conflicts of Interest

Author Haifeng Wang and Ximei Tang were employed by the company Sinopec Petroleum Engineering Corporation. The remaining authors declare that the research was conducted in the absence of any commercial or financial relationships that could be construed as a potential conflict of interest.

References

  1. Mi, Y.; Chen, H.; Li, X.; Meng, Z.; Yang, Q.; Su, H.; Zhou, M.; Wei, Y.; Wu, F. Arbuscular mycorrhizal fungi distribution responds to ecological damage characteristics in antimony mining ecosystems. J. Environ. Manag. 2025, 393, 127194. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  2. Du, J.; Wang, M.; Gao, X.; Wang, X.; Zhang, J.; Fan, H.; Lv, J.; Huang, X. Effects of antidepressant exposures on a stream detrital food chain: Microbial decomposers and invertebrate detritivores. Ecotoxicol. Environ. Saf. 2025, 303, 118974. [Google Scholar] [CrossRef] [Scilit]
  3. Ma, X.; Zhou, Z.; Chen, J.; Xu, H.; Ma, S.; Dippold, M.A.; Kuzyakov, Y. Long-term nitrogen and phosphorus fertilization reveals that phosphorus limitation shapes the microbial community composition and functions in tropical montane forest soil. Sci. Total Environ. 2023, 854, 158709. [Google Scholar] [CrossRef] [Scilit]
  4. Atreya, M.; Desousa, S.; Kauzya, J.B.; Williams, E.; Hayes, A.; Dikshit, K.; Nielson, J.; Palmgren, A.; Khorchidian, S.; Liu, S.; et al. A Transient Printed Soil Decomposition Sensor Based on a Biopolymer Composite Conductor. Adv. Sci. 2023, 10, e2205785. [Google Scholar] [CrossRef] [Scilit]
  5. Moravcova, A.; Barbi, F.; Algora, C.; Tosadori, G.; Macek, P.; Albrechtova, J.; Baldrian, P.; Kohout, P. Contrasting stability of fungal and bacterial communities during long-term decomposition of fungal necromass in Arctic tundra. Environ. Microbiome 2025, 20, 75. [Google Scholar] [CrossRef] [Scilit]
  6. Du, S.; Li, X.Q.; Feng, J.; Huang, Q.; Liu, Y.R. Soil core microbiota drive community resistance to mercury stress and maintain functional stability. Sci. Total Environ. 2023, 894, 165056. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  7. Wang, L.; Ren, X.; Wang, X.; Ye, P.; Wang, F.; Cheng, J.; Chen, Y.; Yu, A.; Zhang, L.; Qiu, Y. Polycyclic aromatic hydrocarbons (PAHs) in the upstream rivers of Taihu Lake Basin, China: Spatial distribution, sources and environmental risk. Environ. Sci. Pollut. Res. Int. 2022, 29, 23690–23699. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  8. Zhao, Y.; Li, J.; Qi, Y.; Guan, X.; Zhao, C.; Wang, H.; Zhu, S.; Fu, G.; Zhu, J.; He, J. Distribution, sources, and ecological risk assessment of polycyclic aromatic hydrocarbons (PAHs) in the tidal creek water of coastal tidal flats in the Yellow River Delta, China. Mar. Pollut. Bull. 2021, 173, 113110. [Google Scholar] [CrossRef] [Scilit]
  9. Guo, W.; Luo, X.; Hou, G.; Liu, F.; Yin, Y.; Li, J.; Shi, X. A century-long record of polycyclic aromatic hydrocarbon deposition in the Old Yellow River Estuary, China. Mar. Pollut. Bull. 2023, 196, 115643. [Google Scholar] [CrossRef] [Scilit]
  10. Gao, Y.; Yuan, L.; Du, J.; Wang, H.; Yang, X.; Duan, L.; Zheng, L.; Bahar, M.; Zhao, Q.; Zhang, W.; et al. Bacterial community profile of the crude oil-contaminated saline soil in the Yellow River Delta Natural Reserve, China. Chemosphere 2022, 289, 133207. [Google Scholar] [CrossRef] [Scilit]
  11. Shaoping, K.; Zhiwei, D.; Bingchen, W.; Huihui, W.; Jialiang, L.; Hongbo, S. Changes of sensitive microbial community in oil polluted soil in the coastal area in Shandong, China for ecorestoration. Ecotoxicol. Environ. Saf. 2021, 207, 111551. [Google Scholar] [CrossRef] [Scilit]
  12. Wang, A.; Fu, W.; Feng, Y.; Liu, Z.; Song, D. Synergetic effects of microbial-phytoremediation reshape microbial communities and improve degradation of petroleum contaminants. J. Hazard. Mater. 2022, 429, 128396. [Google Scholar] [CrossRef] [Scilit]
  13. Lu, W.; Zheng, Y.; Wang, Y.; Song, J.; Weng, Y.; Ma, W.; Arslan, M.; El-Din, M.G.; Wang, D.; Wang, Q.; et al. Survival strategies and assembly mechanisms of microbial communities in petroleum-contaminated soils. Environ. Res. 2024, 262, 119857. [Google Scholar] [CrossRef] [Scilit]
  14. Li, Y.; Li, C.; Xin, Y.; Huang, T.; Liu, J. Petroleum pollution affects soil chemistry and reshapes the diversity and networks of microbial communities. Ecotoxicol. Environ. Saf. 2022, 246, 114129. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  15. Sun, Y.; Ding, A.; Zhao, X.; Chang, W.; Ren, L.; Zhao, Y.; Song, Z.; Hao, D.; Liu, Y.; Jin, N.; et al. Response of soil microbial communities to petroleum hydrocarbons at a multi-contaminated industrial site in Lanzhou, China. Chemosphere 2022, 306, 135559. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  16. Hu, X.; Pan, Y.; Zhao, S.; Zhang, F.; Li, F.; Li, Z.; Li, H.; Li, Y.; Lu, J.; Yang, X.; et al. A step closer to real practice: Integrated tandem photocatalysis-biofilm process towards degradation of crude oil. J. Environ. Manag. 2023, 342, 118357. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  17. Xiao, W.; Lin, G.; He, X.; Yang, Z.; Wang, L. Interactions among heavy metal bioaccessibility, soil properties and microbial community in phyto-remediated soils nearby an abandoned realgar mine. Chemosphere 2022, 286, 131638. [Google Scholar] [CrossRef] [Scilit]
  18. Chi, Z.; Hou, L.; Li, H.; Wu, H.; Yan, B. Indigenous bacterial community and function in phenanthrene-polluted coastal wetlands: Potential for phenanthrene degradation and relation with soil properties. Environ. Res. 2021, 199, 111357. [Google Scholar] [CrossRef] [Scilit]
  19. Louisson, Z.; Hermans, S.M.; Buckley, H.L.; Case, B.S.; Taylor, M.; Curran-Cournane, F.; Lear, G. Land use modification causes slow, but predictable, change in soil microbial community composition and functional potential. Environ. Microbiome 2023, 18, 30. [Google Scholar] [CrossRef] [Scilit]
  20. Saibu, S.; Adebusoye, S.A.; Oyetibo, G.O. Soil microbiome response to 2-chlorodibenzo-p-dioxin during bioremediation of contaminated tropical soil in a microcosm-based study. J. Hazard. Mater. 2023, 451, 131105. [Google Scholar] [CrossRef] [Scilit]
  21. Gao, H.; Wu, M.; Liu, H.; Xu, Y.; Liu, Z. Effect of petroleum hydrocarbon pollution levels on the soil microecosystem and ecological function. Environ. Pollut. 2022, 293, 118511. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  22. Huang, L.; Ye, J.; Jiang, K.; Wang, Y.; Li, Y. Oil contamination drives the transformation of soil microbial communities: Co-occurrence pattern, metabolic enzymes and culturable hydrocarbon-degrading bacteria. Ecotoxicol. Environ. Saf. 2021, 225, 112740. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  23. Liu, H.; Wu, M.; Guo, X.; Gao, H.; Xu, Y. Isotope fractionation (δ13C, δ15N) and microbial community response in degradation of petroleum hydrocarbons by biostimulation in contaminated soil. Environ. Sci. Pollut. Res. Int. 2022, 29, 7604–7613. [Google Scholar] [CrossRef] [Scilit]
  24. Chen, X.; Luo, M.; Tan, J.; Zhang, C.; Liu, Y.; Huang, J.; Tan, Y.; Xiao, L.; Xu, Z. Salt-tolerant plant moderates the effect of salinity on soil organic carbon mineralization in a subtropical tidal wetland. Sci. Total Environ. 2022, 837, 155855. [Google Scholar] [CrossRef] [Scilit] [PubMed]
  25. Liu, Q.; Chen, H.; Su, Y.; Sun, S.; Zhao, C.; Zhang, X.; Gu, Y.; Li, L. Enhanced crude oil degradation by remodeling of crude oil-contaminated soil microbial community structure using sodium alginate/graphene oxide/Bacillus C5 immobilized pellets. Environ. Res. 2023, 223, 115465. [Google Scholar] [CrossRef] [Scilit]
  26. Shi, H.; Gao, W.; Zheng, Y.; Yang, L.; Han, B.; Zhang, Y.; Zheng, L. Distribution and abundance of oil-degrading bacteria in seawater of the Yellow Sea and Bohai Sea, China. Sci. Total Environ. 2023, 902, 166038. [Google Scholar] [CrossRef] [Scilit]
  27. Raiyani, N.M.; Singh, S.P. Microbial community and predictive functionalities associated with the marine sediment of Coastal Gujarat. Environ. Sci. Pollut. Res. Int. 2023, 30, 43245–43266. [Google Scholar] [CrossRef] [Scilit]
  28. Wan, P.; Zhou, Z.; Yuan, Z.; Wei, H.; Huang, F.; Li, Z.; Li, F.-M.; Zhang, F. Fungal community composition changes and reduced bacterial diversity drive improvements in the soil quality index during arable land restoration. Environ. Res. 2024, 244, 117931. [Google Scholar] [CrossRef] [Scilit]
  29. Zhu, N.; Wang, J.; Wang, Y.; Li, S.; Chen, J. Differences in geological conditions have reshaped the structure and diversity of microbial communities in oily soils. Environ. Pollut. 2022, 306, 119404. [Google Scholar] [CrossRef] [Scilit]
Figure 1. Sampling site.
Diversity 18 00206 g001
Figure 2. Rarefaction curves of all 30 samples. The vertical dashed line indicates the rarefaction depth of 30,000 sequences per sample used for diversity analyses.
Diversity 18 00206 g002
Figure 3. ASV cluster analysis.
Diversity 18 00206 g003
Figure 4. Analysis of microbial composition characteristics among two groups. (A): The composition of microorganisms at the family level; (B): The composition of microorganisms at the genus level.
Diversity 18 00206 g004
Figure 5. α-Diversity analysis of the microbiota among two groups.
Diversity 18 00206 g005
Figure 6. β-diversity analysis of the microbiota among two groups. (A): the PCoA analysis between two groups; (B): Analysis of inter-group differences between the two groups.
Diversity 18 00206 g006
Figure 7. LEfSe analysis among two groups.
Diversity 18 00206 g007
Figure 8. Functional predictive analysis among two groups. (A): PICRUSt2 analysis among two groups; (B): FAPROTAX-based analysis among two groups.
Diversity 18 00206 g008
Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content.