


Abstract
β-Lactosylceramide (β-LacCer) is not only a key intermediate in the biosynthesis of complex glycosphingolipids (GSLs) but also an important regulator of many biological processes. To facilitate the investigation of β-LacCer and other GSLs, a series of β-LacCer analogs with an azido group at the 6-C-position of the D-galactose in lactose and varied forms of the ceramide moiety were synthesized from commercially available lactose in sixteen linear steps by a versatile and diversity-oriented strategy, which engaged lipid remodeling and glycan functionalization at the final stage. These azide-labeled β-LacCer analogs are flexible and universal platforms that are suitable for further functionalization with other molecular tags via straightforward and biocompatible click chemistry, thereby paving the way for their application to various biological studies.
Keywords:
carbohydrates; glycolipids; glycosphingolipids; β-Lactosylceramide; lipids; lactose; azide; synthesis 1. Introduction
Glycosphingolipids (GSLs) are ubiquitous and abundant glycolipids in the cell membrane [1] that are composed of a hydrophilic glycan head and a lipophilic ceramide (Cer) tail coupled together by a glycosidic bond [2,3]. The amphiphilic property of GSLs enables them to adhere to the cell membrane outer leaflet and traffic through the lipid bilayer [4,5]. Inside the cell membrane, GSLs tend to self-aggregate in specific microdomains of the lipid rafts, where signaling molecules are localized and enriched [6,7]. As a result, GSLs play an important role in numerous biophysiological processes, such as cellular recognition and signaling, cell differentiation and proliferation, etc. [8]. GSL dysregulation is also associated with many human diseases, including cancer and Alzheimer’s disease [9,10]. For example, a number of GSLs have been identified as tumor-associated carbohydrate antigens (TACAs) and utilized as molecular targets for the development of therapeutic cancer vaccines [9,11].
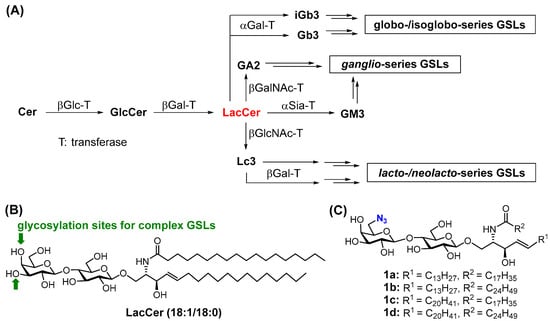
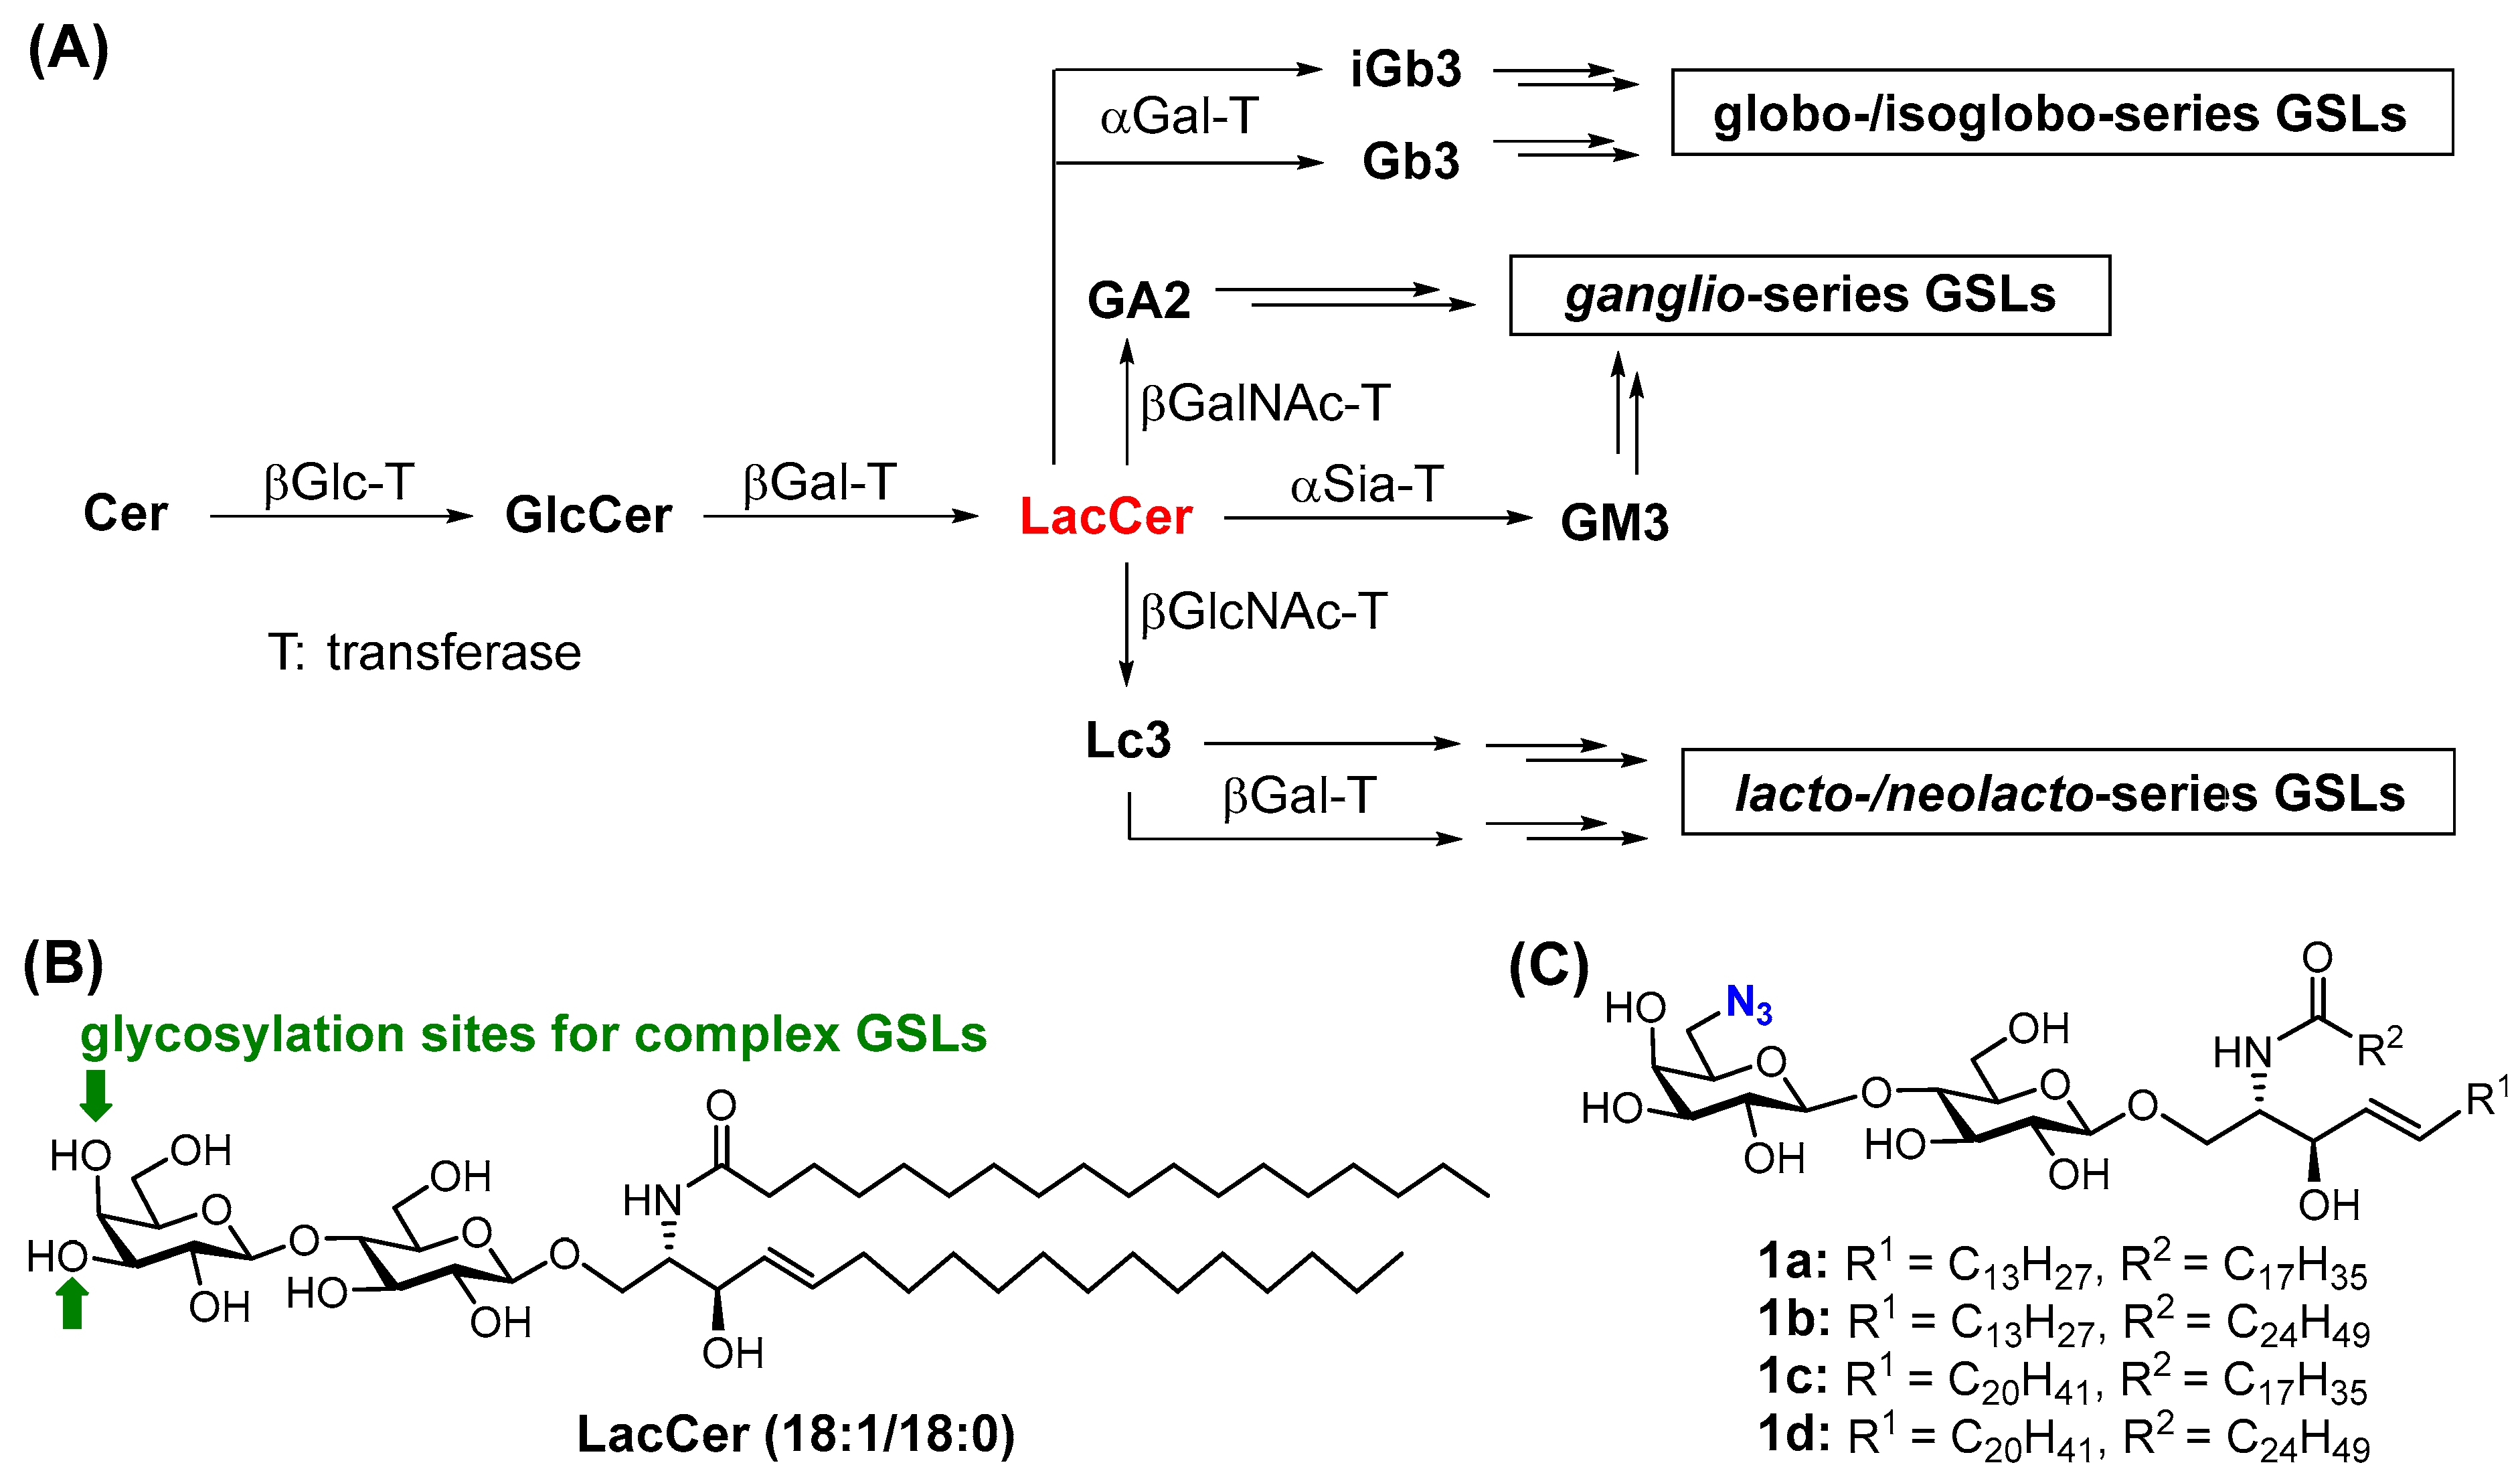
Natural GSLs are structurally diverse, with potential structural variations in both the glycan and the Cer moiety [1]. β-Lactosylceramide (β-LacCer) is a simple GSL having the D-glucose (Glc) residue of lactose, a disaccharide, β-linked to the Cer moiety. In mammals, it is the key intermediate during the biosynthesis of the majority of complex GSLs, e.g., globo-, isoglobo-, ganglio-, lacto-, and neolecto-series GSLs (Figure 1A) [12,13]. In the meantime, it has been demonstrated that β-LacCer is involved in the human central nervous system, immune system, and diseases like cancer, inflammation, and neurodegeneration [14,15,16]. For instance, β-LacCer overexpression is found in leukemia, renal cancer, and cholangiocarcinoma [17], and LacCer accumulation is also related to atherosclerosis and aberrant autophagy [18,19]. Thus, β-LacCer is identified as a drug target [20]. Additionally, similar to other GSLs, the lipids in β-LacCer usually contain 16–20 carbons (C16–20) (such as C18, Figure 1B), while the forms of β-LacCer with longer (>C24) lipid chains are also found in neutrophils, which become molecular targets for the treatment of immunological disorders [21,22,23].
Figure 1.
(A) The biosynthetic pathways for globo-, isoglobo-, ganglio-, lacto-, and neolecto-series GSLs, utilizing β-LacCer as the key intermediate, and some key enzymes involved, and structures of (B) a representative β-LacCer with the most common (18:1/18:0) lipid form found in mammals and (C) designed β-LacCer analogs with different lipid forms as useful probes to study β-LacCer and other GSLs.
However, detailed investigation of β-LacCer, especially within complex matrices like the lipid membrane, is challenging due to the lack of fluorophores and other visible functionalities in its structure. To address the problem, it is necessary to functionalize β-LacCer with molecular tags (e.g., affinity and fluorescent tags) to facilitate various modern analytical technologies. In this context, we designed a series of β-LacCer analogs 1a–d (Figure 1C) containing an azido group and diverse lipid chains and synthesized them by a diversity-oriented strategy. These β-LacCer analogs should be useful molecular tools.
2. Results and Discussion
In the designed β-LacCer probes 1a–d (Figure 1C), we planned to attach an azido group to the 6-C-position of D-galactose (Gal) in the lactose moiety for several reasons. First, the azido group is a flexible platform to facilitate further functionalization of 1a–d with various molecular labels through straightforward and biocompatible click chemistry [24,25]. Next, the azido group is small—not significantly larger than a hydroxyl group, and it is expected to have a minimal impact on the properties of β-LacCer. Consequently, if 1a–d are used as biosynthetic precursors for metabolic glycoengineering, they are likely to be acceptable by enzymes. Moreover, the 6-C-position of the Gal residue in lactose is distinct, because it is a primary carbon at the non-reducing end and is probably the least sterically hindered position, making it relatively efficient and straightforward to functionalize this position and subsequently modify the synthetic probes. The designed probes also contain different forms of lipids, which will facilitate the investigation of how lipid structures of β-LacCer influence its chemical, biophysical, and biological properties.
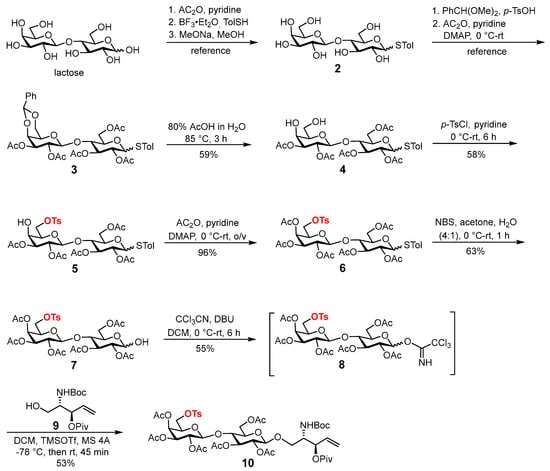
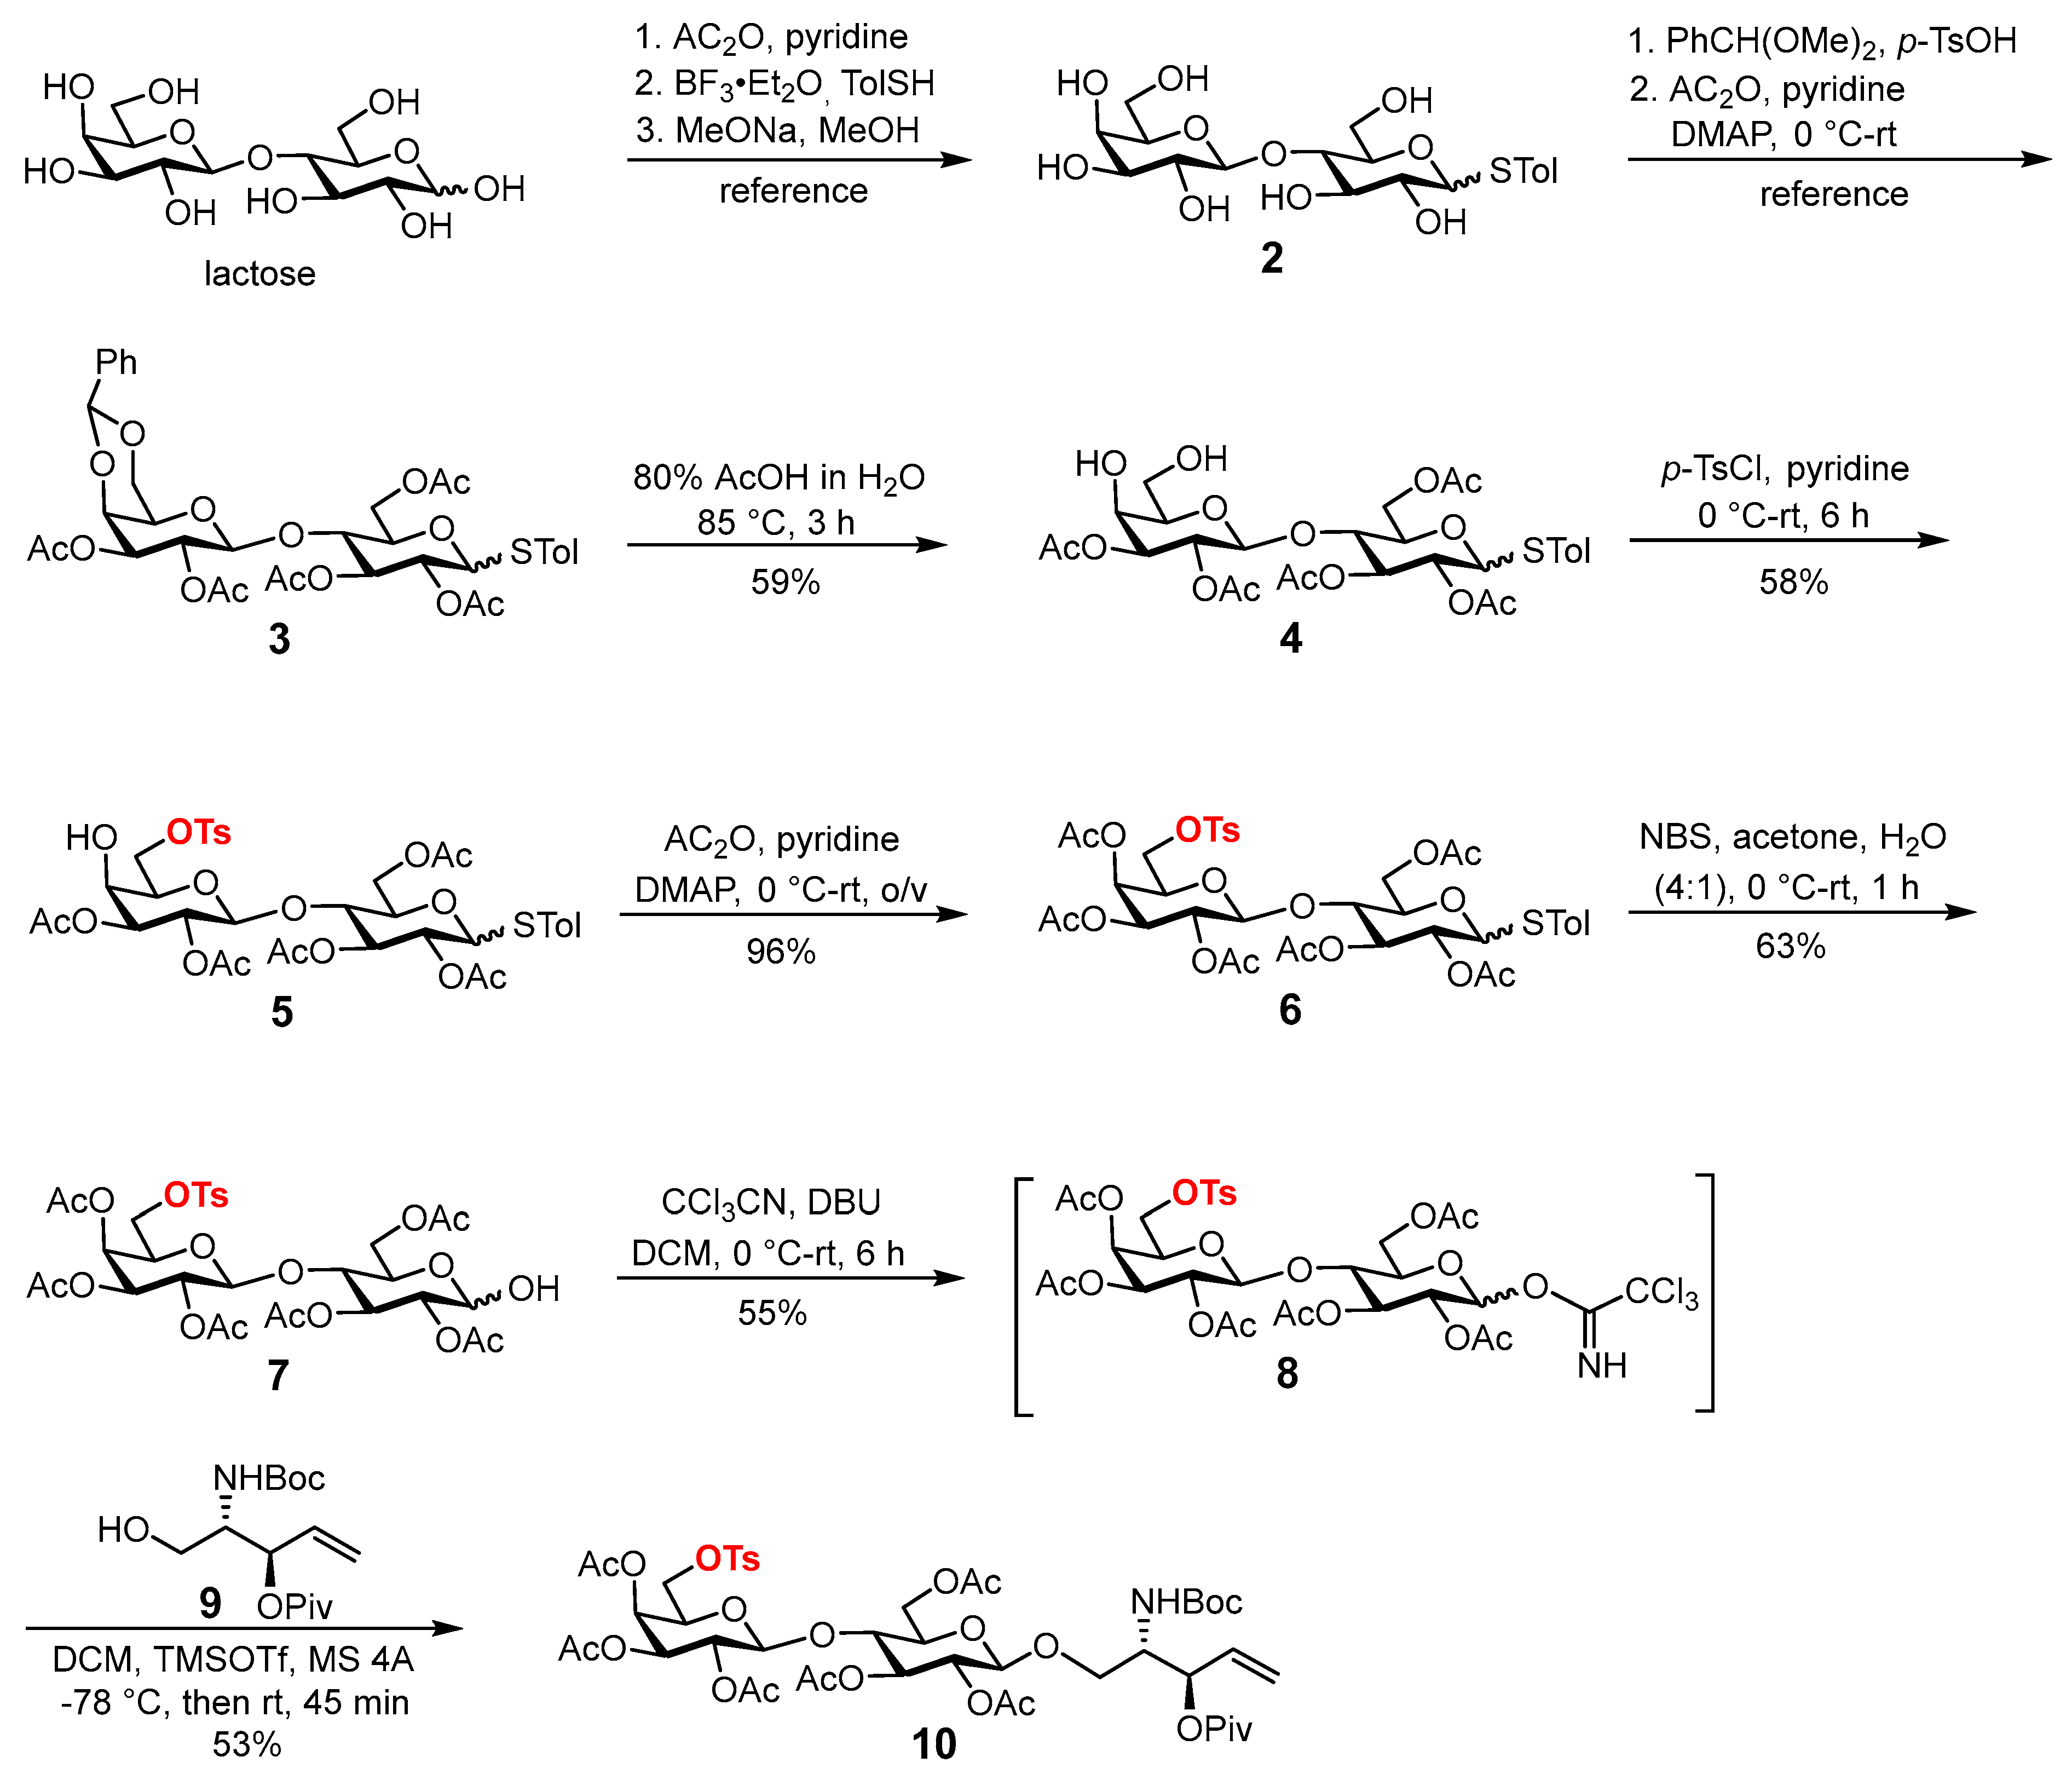
Our synthesis of the target molecules 1a–d (Scheme 1) started with commercially available and inexpensive lactose. It was first converted into the O-acetyl and 4′,6′-O-benzylidene protected glycoside of p-toluenethiol (TolSH) 3 as an α,β-isomeric mixture via a series of well-established reactions reported in the literature [26,27,28,29]. Next, the 4′,6′-O-benzylidene group in 3 was selectively removed upon treatment with 80% acetic acid in water at 85 °C to afford diol 4 as an anomeric mixture. Selective tosylation of the 6′-OH group in 4 was achieved by its reaction with p-toluenesulfonylchloride (p-TsCl) and 4-dimethylaminopyridine (DMAP). The regioselective attack of p-TsCl to the 6′-OH group is probably because this primary hydroxyl group is less sterically hindered than the secondary 4′-OH group in 4. The equivalence and concentration of p-TsCl, along with the order to add reagents, are also substantial in the control of this regioselectivity. The formation of 5 was confirmed by the downfield shifts of its 6′-H NMR signals, compared to those of 4 (from δ 4.06 and 3.72 to 4.41 and 4.07 ppm), and the appearance of 1H signals of an additional toluene moiety at δ 7.40–7.35 and 2.47 ppm, respectively, in the 1H NMR spectrum of 5. The free hydroxyl group remaining in 5 was then acetylated using acetic anhydride, pyridine, and DMAP. To reveal the reducing-end anomeric position for glycosidation, the resultant 6 was applied to oxidative hydrolysis with N-bromosuccinimide (NBS) in water and acetone (1:4) to afford hemiacetal 7 (63%) as an anomeric mixture, together with unreacted 6 (34%). Next, 7 was converted into imidate 8 as a glycosyl donor upon reacting with trichloroacetonitrile catalyzed by 1,8-diazabicyclo-[5.4.0]-undec-7-ene (DBU). After brief purification, the reactive glycosyl imidate 8 was immediately submitted to the glycosylation reaction with Cer precursor 9 [30,31], utilizing trimethylsilyltrifluoromethanesulfonate (TMSOTf) as the promotor. This reaction produced 10 as the key synthetic intermediate in excellent stereoselectivity, and the β-configuration of its newly formed glycosidic bond is confirmed by the large coupling constant (J = 7.8 Hz) of its anomeric proton at δ 4.43 in the 1H NMR spectrum. The common intermediate 10 was then used for diversity-oriented synthesis of all β-LacCer probes 1a–d.
Scheme 1.
Preparation of 10—the key and common building block for the synthesis of all designed probes in the present work.
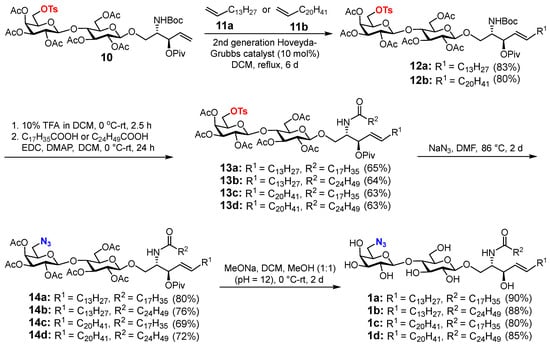
The final steps for 1a–d synthesis include lipid remodeling, glycan functionalization, and global deprotection, as outlined in Scheme 2. Olefin cross-metathesis of pentadecene 11a or docosene 11b with 10 in the presence of the 2nd generation Hoveyda−Grubbs catalyst [32] allowed on-site generation of the sphingosine moiety of varied chain lengths in very good yields (80–83%), although the reaction was rather slow, taking six days to complete [31]. The E-configuration of the C=C bond in 12a and 12b was verified by the olefinic 1H-1H coupling constant (J = 14.6–14.8 Hz) in their 1H NMR spectra. To attach the N-fatty acyl chain, the tert-butoxycarbonyl (Boc) group in 12a,b was selectively removed with 10% trifluoroacetic acid (TFA) in dichloromethane (DCM) to afford the intermediate free amines, which were directly subjected to N-acylation employing stearic acid or pentacosanoic acid, with N-(3-dimethylaminopropyl)-N′-ethylcarbodiimide hydrochloride (EDC) and DMAP as the condensation reagents. These reactions gave 6′-O-tosylated β-LacCer derivatives 13a–d containing different lipid chains in good overall yields (63–65%, two steps). Subsequently, nucleophilic substitution of the tosylate in 13a–d with sodium azide in dimethylformamide (DMF) proceeded smoothly to provide fully protected LacCer analogs 14a–d in high yields (69–80%). Eventually, all O-acyl groups in 14a–d were removed with sodium methoxide in methanol and DCM (1:1) to produce the synthetic targets 1a–d in 80–90% yields. The final products, as well as all new intermediates involved in these syntheses, have been fully characterized with NMR and high-resolution mass spectrometry (HR-MS) data (see data and Figures in Supporting Information).
Scheme 2.
Assembly of the designed β-LacCer probes 1a–d.
3. Experimental Section
3.1. General Methods
Chemicals, reagents, and solvents were commercial and used without further purification. Molecular sieves 4 Å (MS 4Å) were flame-dried under a high vacuum and used after being cooled to room temperature (rt) under an N2 atmosphere. Thin-layer chromatography (TLC) was performed on silica gel 60 Å F254 plates, with detection utilizing a UV lamp at a wavelength of 254 nm followed by charring with 10% (v/v) H2SO4 in ethanol or ninhydrin and anisaldehyde staining. Flash column chromatography was performed using silica gel 60 (230–400 mesh). NMR spectra were acquired on a 400, 500, or 600 MHz spectrometer, with chemical shifts (δ) reported in ppm as referenced to internal tetramethylsilane (TMS) (1H NMR: δ 0.00 ppm), CDCl3 (1H NMR: δ 7.26 ppm; 13C{1H} NMR: δ 77.2 ppm), or CD3OD (1H NMR: δ 3.31 ppm; 13C{1H} NMR: δ 49.0 ppm). Coupling constants (J) were reported in Hz. Peak, and coupling constant assignments were based upon 1H NMR, 1H–1H COSY, and 1H–13C{1H} HSQC experiments. HR-MS spectra were recorded on the XEVO-G2-XS Q-TOF-ESI instrument. An aluminum heating block was used for heating reaction mixtures. Intermediates 2, 3, and 9 were prepared according to the reported protocols, and their 1H NMR spectra were the same as those in the literature [26,27,28,29].
3.2. p-Methylphenyl 2,3-di-O-acetyl-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-1-thio-β-D-glucopyranoside (4)
A solution of 3 (51.5 g, 68.96 mmol) in 80% aqueous AcOH was heated at 85 °C using an aluminum block for 3 h. After the reaction was complete, as indicated by the consumption of all starting material on TLC, the solvent was removed under reduced pressure followed by co-evaporation with toluene three times. Thereafter, ethyl acetate (EtOAc) was added, and the solution was washed with NaHCO3, water, and brine multiple times. The organic layer was dried over MgSO4, filtered, and concentrated under vacuum. The crude product was purified using silica gel column chromatography to afford 4 (59%, 26.8 g) as a white solid. TLC: Rf = 0.61 (EtOAc:Hex 80:20). 1H NMR (500 MHz, CDCl3): δ 7.35 (d, J = 8.0 Hz, 2H), 7.09 (d, J = 8.0 Hz, 2H), 5.31–5.02 (m, 2H), 4.95–4.79 (m, 2H), 4.61 (d, J = 10.1 Hz, 1H, anomeric), 4.53 (dd, J = 11.7, 1.9 Hz, 1H), 4.46 (d, J = 7.9 Hz, 1H, anomeric), 4.08 (dt, J = 11.9, 8.5 Hz, 2H), 3.93–3.72 (m, 3H), 3.68–3.48 (m, 2H), 3.45–3.30 (m, 1H), 2.79 (dd, J = 7.1, 4.7 Hz, 1H), 2.33 (s, 3H), 2.09 (s, 3H), 2.07 (s, 3H), 2.05 (s, 6H, 2 × CH3), 2.02 (s, 3H). 13C NMR (126 MHz, CDCl3): δ 170.8, 170.6, 170.4, 169.71, 169.7, 138.7, 133.7, 129.7, 127.8, 101.1, 85.6, 76.7, 76.3, 74.5, 73.6, 70.4, 70.3, 69.8, 67.8, 62.4, 62.1, 21.3, 21.01, 21.0, 20.9, 20.8. HR-ESI-MS m/z: [M + H2O + H]+ Calcd for C29H41O16S 677.2115; Found 677.2106.
3.3. p-Methylphenyl 2,3-di-O-acetyl-6-O-(p-toluenesulfonyl)-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-1-thio-β-D-glucopyranoside (5)
To a solution of 4 (20.5 g, 31.12 mmol) dissolved in pyridine (93.2 mL) was added p-TsCl (5.93 g, 31.12 mmol) portionwise under an N2 atmosphere in an ice bath. The mixture was stirred at rt for 6 h, at which point the reaction did not show further progress. Next, MeOH was added dropwise to the reaction mixture, and the solvent was removed under vacuo. The residue was extracted with EtOAc and washed twice with brine. The organic layer was dried over MgSO4 and concentrated under reduced pressure. Purification of the product by silica gel column chromatography afforded 5 (58%, 14.67 g) as a white waxy solid. TLC: Rf = 0.95 (EtOAc:Hex 80:20). 1H NMR (400 MHz, CDCl3): δ 7.78 (d, J = 8.3 Hz, 2H), 7.40–7.35 (m, 4H), 7.10 (d, J = 8.0 Hz, 2H), 5.15 (t, J = 9.1 Hz, 1H), 5.09 (dd, J = 10.2, 7.9 Hz, 1H), 4.92–4.86 (m, 1H), 4.83 (d, J = 9.4 Hz, 1H), 4.58 (d, J = 10.1 Hz, 1H, anomeric), 4.50 (dd, J = 11.9, 1.9 Hz, 1H), 4.41 (d, J = 7.9 Hz, 1H, anomeric), 4.24 (dd, J = 10.4, 6.5 Hz, 1H), 4.12–4.05 (m, 2H), 4.03 (t, J = 4.4 Hz, 1H), 3.76 (t, J = 6.4 Hz, 1H), 3.68 (t, J = 9.5 Hz, 1H), 3.58 (ddd, J = 10.0, 5.3, 1.9 Hz, 1H), 2.47 (s, 3H), 2.33 (s, 3H), 2.10 (s, 3H), 2.08 (s, 3H), 2.06 (s, 3H), 2.01 (s, 3H), 1.90 (s, 3H). 13C NMR (101 MHz, CDCl3): δ 170.5, 170.2, 170.1, 169.6, 169.5, 145.5, 138.7, 133.7, 132.6, 130.2, 129.7, 128.0, 127.9, 100.8, 85.6, 76.7, 75.9, 74.0, 73.2, 72.3, 70.3, 69.5, 67.1, 66.5, 62.2, 21.8, 21.3, 20.92, 20.91, 20.9, 20.8, 20.7. HR-ESI-MS m/z: [M + NH4]+ Calcd for C36H48NO17S2 830.2364; Found 830.2354
3.4. p-Methylphenyl 2,3,4-tri-O-acetyl-6-O-(p-toluenesulfonyl)-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-1-thio-β-D-glucopyranoside (6)
After 5 (14.5 g, 17.84 mmol) was dissolved in pyridine (90 mL), Ac2O (2.53 mL, 26.76 mmol) and DMAP (0.4 g, 3.57 mmol) were added at 0 °C under an N2 atmosphere. The mixture was stirred at rt overnight. After the complete consumption of 5 as indicated by TLC, MeOH was added dropwise to quench the reaction. The solvent was evaporated under vacuo. The resulting residue was extracted with EtOAc, and the organic layer was washed three times with H2O and brine, dried over MgSO4, and finally, concentrated under vaccum. The residue was purified by silica gel column chromatography to afford 6 (96%, 14.6 g) as a white waxy solid. TLC: Rf = 0.66 (DCM:MeOH 95:5). 1H NMR (400 MHz, CDCl3): δ 7.75 (d, J = 8.3 Hz, 2H), 7.36 (dd, J = 8.1, 1.7 Hz, 4H), 7.10 (d, J = 7.9 Hz, 2H), 5.32 (dd, J = 3.4, 0.8 Hz, 1H), 5.18 (t, J = 9.1 Hz, 1H), 5.04 (dd, J = 10.4, 7.9 Hz, 1H), 4.91 (dd, J = 10.4, 3.4 Hz, 1H), 4.83 (t, J = 9.6 Hz, 1H), 4.59 (d, J = 10.1 Hz, 1H, anomeric), 4.50 (dd, J = 11.9, 2.0 Hz, 1H), 4.44 (d, J = 7.9 Hz, 1H, anomeric), 4.08–3.95 (m, 3H), 3.88 (dd, J = 7.5, 6.5 Hz, 1H), 3.70 (t, J = 9.5 Hz, 1H), 3.59 (ddd, J = 10.0, 5.2, 1.9 Hz, 1H), 2.46 (s, 3H), 2.33 (s, 3H), 2.10 (s, 3H), 2.09 (s, 3H), 2.04 (s, 3H), 2.01 (s, 3H), 1.95 (s, 3H), 1.94 (s, 3H). 13C NMR (101 MHz, CDCl3): δ 170.4, 170.01, 170.0, 169.8, 169.6, 169.1, 145.6, 138.8, 133.8, 132.3, 130.2, 129.8, 128.1, 127.8, 100.9, 85.6, 76.7, 76.1, 74.0, 70.91, 70.9, 70.4, 69.1, 66.6, 65.5, 62.1, 21.8, 21.3, 21.01, 21.0, 20.9, 20.7, 20.61, 20.6. HR-ESI-MS m/z: [M + HCOO]− Calcd for C39H47O20S2 899.2103; Found 899.2074.
3.5. 2,3,4-Tri-O-acetyl-6-O-(p-toluenesulfonyl)-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-D-glucopyranose (7)
To a solution of 6 (8.0 g, 9.36 mmol) dissolved in acetone (160 mL) and H2O (40 mL) was added NBS (3.33 g, 18.72 mmol) at 0 °C. The mixture was stirred at rt for 1 h, while the reaction was monitored by TLC. Once no further progress was observed with TLC, the reaction was quenched by adding solutions of Na2S2O3 and NaHCO3 dropwise. Acetone was removed under vacuum and then saturated aq. NaHCO3 solution was added to the residue. The mixture was extracted with DCM three times. The organic layers were combined, dried over MgSO4, and concentrated under vacuum. The residue was purified by silica gel column chromatography to afford 7 (63%, 4.40 g) as a white solid, along with recovery of 6 (34%, 2.70 g). TLC: Rf = 0.25 (EtOAc:Hex 60:40). 1H NMR (400 MHz, CDCl3): δ 7.77 (d, J = 8.3 Hz, 2H), 7.37 (d, J = 8.4 Hz, 2H), 5.54 (t, J = 9.7 Hz, 1H), 5.38 (t, J = 3.4 Hz, 1H), 5.32 (d, J = 2.5 Hz, 1H), 5.09 (dd, J = 10.4, 7.9 Hz, 1H), 4.92 (dd, J = 10.4, 3.4 Hz, 1H), 4.83 (dd, J = 10.1, 3.4 Hz, 1H, anomeric), 4.53 (d, J = 7.9 Hz, 1H, anomeric), 4.48 (ddd, J = 10.4, 7.3, 2.3 Hz, 2H), 4.23 (ddd, J = 10.1, 4.1, 1.9 Hz, 1H), 4.12 (dd, J = 12.0, 4.1 Hz, 1H), 4.07 (dd, J = 10.6, 4.2 Hz, 2H), 3.96 (t, J = 3.3 Hz, 1H), 3.94–3.90 (m, 1H), 3.79 (t, J = 9.7 Hz, 1H), 2.13 (s, 3H), 2.09 (s, 3H), 2.06 (s, 3H), 2.03 (s, 3H), 2.01 (s, 3H), 1.94 (s, 3H). 13C NMR (101 MHz, CDCl3): δ 170.7, 170.4, 170.1, 169.8, 169.2, 169.1, 132.3, 130.2, 129.8, 128.2, 100.6, 90.3, 77.4, 76.3, 71.6, 71.0, 69.3, 69.1, 68.3, 66.8, 66.0, 62.0, 21.8, 21.1, 21.0, 20.9, 20.8, 20.7, 20.6. HR-ESI-MS m/z: [M-H2O + H]+ Calcd for C31H39O18S 731.1857; Found: 731.1833.
3.6. 2,3,4-Tri-O-acetyl-6-O-(p-toluenesulfonyl)-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-D-glucopyranosyl trichloroimidate (8)
To a solution of 7 (990 mg, 1.322 mmol) in dry DCM (9.5 mL) were added DBU (0.132 mmol, 20 µL) and trichloroacetonitrile (2.12 mL, 21.15 mmol) dropwise at 0 °C under an N2 atmosphere. The mixture was stirred at rt for 3 h. After the reaction was completed as indicated by TLC, the solvent was removed under reduced pressure, and the residue was purified by flash column chromatography using Et3N-deactivated silica to give 8 (55%, 649 mg). TLC: Rf = 0.45 (EtOAc:Hex 50:50). 1H NMR (400 MHz, CDCl3): δ 8.66 (s, 1H, NH), 7.77 (d, J = 8.2 Hz, 2H), 7.38 (d, J = 8.1 Hz, 2H), 6.49 (d, J = 3.7 Hz, 1H, anomeric), 5.55 (t, J = 9.6 Hz, 1H), 5.33 (d, J = 3.3 Hz, 1H), 5.13–5.08 (m, 1H), 5.05 (dd, J = 10.2, 3.9 Hz, 1H), 4.93 (dd, J = 10.4, 3.4 Hz, 1H), 4.51 (d, J = 8.0 Hz, 1H, anomeric), 4.49–4.44 (m, 1H), 4.14–4.05 (m, 1H), 3.99 (dd, J = 10.0, 6.6 Hz, 1H), 3.95–3.81 (m, 1H), 2.11 (s, 3H), 2.06 (s, 3H), 2.03 (s, 6H, 2 × CH3), 2.02 (s, 3H), 1.95 (s, 3H). 13C NMR (101 MHz, CDCl3): δ 170.4, 170.2, 170.1, 170.0, 169.5, 169.2, 161.1, 145.7, 132.3, 130.2, 128.2, 101.3, 93.0, 77.4, 76.0, 71.1, 71.0, 70.2, 69.7, 69.1, 66.7, 65.6, 61.6, 21.8, 21.2, 21.05, 21.0, 20.8, 20.7, 20.6.
3.7. (2S,3R)-2-(tert-Butoxycarbonyl)amino-3-pivaloyloxy-pent-4-en-1-yl 2,3,4-tri-O-acetyl-6-O-(p-toluenesulfonyl)-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-D-glucopyranoside (10)
The mixture of glycosyl donor 8 (600 mg, 0.671 mmol), acceptor 9 (168 mg, 0.559 mmol), and flame-dried MS 4Å (1.5 g) in dry DCM (12 mL) was stirred at rt for 15 min under an N2 atmosphere and then cooled to -78 °C. Thereafter, TMSOTf (4.0 µL, 22.4 µmol) was added dropwise followed by stirring at the same temperature (−78 °C) for 15 min. The mixture was gradually warmed to rt and stirred for another 45 min. When TLS indicated the complete consumption of 9, the reaction was quenched with Et3N at 0 °C, and the mixture was filtered through a Celite pad. The filtrate was washed with saturated NaHCO3solution, and the organic layer was dried over MgSO4 and concentrated under vacuo. Silica gel column chromatography of the residue afforded 10 (53% yield, 306 mg) as a white solid. TLC: Rf = 0.50 (EtOAc:Hex 50:50).1H NMR (600 MHz, CDCl3): δ 7.77 (d, J = 8.3 Hz, 2H), 7.38 (d, J = 8.1 Hz, 2H), 5.83–5.73 (m, 1H, =CH), 5.32 (dd, J = 11.2, 5.1 Hz, 2H), 5.29 (d, J = 1.2 Hz, 1H), 5.26 (d, J = 10.7 Hz, 1H), 5.17 (t, J = 9.3 Hz, 1H), 5.06 (dd, J = 10.3, 7.9 Hz, 1H), 4.92 (dd, J = 10.4, 3.4 Hz, 1H), 4.88 (dd, J = 9.4, 8.0 Hz, 1H), 4.70 (d, J = 8.9 Hz, 1H, -NH), 4.46 (d, J = 7.8 Hz, 1H, anomeric), 4.45–4.44 (m, 1H), 4.43 (d, J = 7.8 Hz, 1H, anomeric), 4.08–3.97 (m, 4H), 3.94 (dd, J = 10.0, 3.4 Hz, 1H), 3.90 (t, J = 6.5 Hz, 1H), 3.77 (t, J = 9.5 Hz, 1H), 3.63–3.57 (m, 1H), 3.51 (dd, J = 10.1, 4.9 Hz, 1H), 2.46 (s, 1H), 2.11 (s, 1H), 2.06 (s, 1H), 2.04 (s, 1H), 2.02 (s, 1H), 1.99 (s, 1H), 1.94 (s, 1H), 1.42 (s, 9H), 1.19 (s, 9H). 13C NMR (151 MHz, CDCl3): δ 177.1, 170.5, 170.01, 170.0, 169.9, 169.8, 169.7, 169.2, 155.3, 145.7, 132.4, 130.2, 128.2, 118.7, 100.91, 100.9, 79.8, 76.1, 73.3, 72.8, 72.6, 71.8, 70.91, 70.9, 69.1, 68.5, 66.7, 65.6, 62.1, 60.5, 52.2, 39.0, 28.5, 27.2, 21.8, 21.2, 21.0, 20.9, 20.71, 20.7, 20.6. HR-ESI-MS m/z: [M + HCOO]− Calcd for C47H66NO25S 1076.3645; Found 1076.3657.
3.8. (2S,3R,E)-2-(tert-Butoxycarbonyl)amino-3-pivaloyloxy-octadec-4-en-1-yl 2,3,4-tri-O-acetyl-6-O-(p-toluenesulfonyl)-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-D-glucopyranoside (12a)
To an N2-bubbled mixture of 10 (150 mg, 0.145 mmol) and 1-pentadecene 11a (183.5 mg, 0.237 mL, 0.872 mmol) in dry DCM (28 mL) was added 2nd generation Hoveyda−Grubbs catalyst (9 mg, 10 mol%). The reaction mixture was refluxed at 40 °C for 6 d. Each day, another batch of 11a (183.5 mg, 0.237 mL, 0.873 mmol) and the catalyst (4.53 mg, 5 mol%) was added to the mixture. Upon the completion of reaction, DMSO (2 drops) was added, and the mixture was stirred at rt for 40 min. The solvent was removed under vacuo, and the residue was purified by silica gel column chromatography to give 12a (83%, 146.5 mg) as a brownish solid. TLC: Rf = 0.83 (EtOAc:Hex 80:20). 1H NMR (400 MHz, CDCl3): δ 7.78 (d, J = 8.3 Hz, 2H), 7.39 (d, J = 8.0 Hz, 2H), 5.75 (dt, J = 14.6, 7.6 Hz, 1H, =CH-), 5.40–5.30 (m, 2H, =CH- and 4′-H), 5.18 (dd, J = 17.4, 8.2 Hz, 2H), 5.06 (dd, J = 10.4, 7.9 Hz, 1H), 4.90 (ddd, J = 17.3, 9.9, 5.7 Hz, 2H), 4.67 (d, J = 9.2 Hz, 1H, NH), 4.46 (d, J = 7.8 Hz, 1H, anomeric), 4.43 (d, J = 7.7 Hz, 1H, anomeric), 4.10–3.96 (m, 4H), 3.91 (dd, J = 13.5, 7.8 Hz, 2H), 3.77 (t, J = 9.4 Hz, 1H), 3.59 (ddd, J = 9.8, 4.7, 1.7 Hz, 1H), 3.49 (dd, J = 9.7, 4.4 Hz, 1H), 2.47 (s, 3H), 2.11 (s, 3H), 2.07 (s, 3H), 2.06 (s, 3H), 2.03 (s, 3H), 1.99 (s, 3H), 1.95 (s, 3H), 1.42 (s, 9H), 1.30–1.21 (m, 24H), 1.17 (s, 9H), 0.88 (t, J = 6.8 Hz, 3H). 13C NMR (101 MHz, CDCl3): δ 177.0, 171.3, 170.5, 170.01, 170.0, 169.9, 169.8, 169.2, 145.6, 137.0, 132.4, 130.2, 128.2, 124.6, 100.91, 100.9, 77.4, 76.1, 73.3, 72.71, 72.7, 71.8, 70.9, 69.1, 68.61, 68.6, 66.6, 65.5, 62.1, 60.5, 52.3, 38.9, 32.5, 32.1, 29.81, 29.8, 29.7, 29.5, 29.3, 29.1, 28.5, 27.2, 22.8, 21.8, 21.2, 21.0, 20.9, 20.8, 20.71, 20.7, 20.6, 14.31, 14.3. HR-ESI-MS m/z: [M + HCOO]− Calcd for C60H92NO25S 1258.5680; found: 1258.5686
3.9. (2S,3R,E)-2-(tert-Butoxycarbonyl)amino-3-pivaloyloxy-pentacos-4-en-1-yl 2,3,4-tri-O-acetyl-6-O-(p-toluenesulfonyl)-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-D-glucopyranoside (12b)
Compound 12b (80%, 152.6 mg) was prepared as a blackish brown solid from 11b by the same method and conditions utilized to prepare 12a. TLC: Rf = 0.92 (EtOAc:Hex 80:20). 1H NMR (600 MHz, CDCl3): δ 7.78 (d, J = 8.3 Hz, 2H), 7.38 (d, J = 8.1 Hz, 2H), 5.75 (dt, J = 14.6, 7.6 Hz, 1H, =CH-), 5.40–5.30 (m, 2H, =CH- and 4′-H), 5.18 (dd, J = 17.3, 8.1 Hz, 2H), 5.06 (dd, J = 10.4, 7.9 Hz, 1H), 4.89 (ddd, J = 17.3, 9.9, 5.7 Hz, 2H), 4.66 (d, J = 8.9 Hz, 1H, NH), 4.46 (d, J = 7.8 Hz, 1H, anomeric), 4.45–4.44 (m, 1H), 4.43 (d, J = 7.6 Hz, 1H, anomeric), 4.05 (dd, J = 11.1, 5.0 Hz, 2H), 3.99 (dd, J = 10.2, 6.6 Hz, 2H), 3.91 (dd, J = 13.4, 7.5 Hz, 2H), 3.77 (t, J = 9.4 Hz, 1H), 3.59 (ddd, J = 9.6, 4.6, 1.5 Hz, 1H), 3.48 (dd, J = 9.7, 4.4 Hz, 1H), 2.47 (s, 3H), 2.10 (s, 3H), 2.06 (s, 6H), 2.02 (s, 3H), 1.99 (s, 3H), 1.94 (s, 3H), 1.42 (s, 9H), 1.36–1.20 (m, 38H), 1.17 (s, 9H), 0.88 (t, J = 6.8 Hz, 3H). 13C NMR (151 MHz, CDCl3): δ 177.0, 170.5, 170.01, 170.0, 169.9, 169.81, 169.8, 169.2, 145.6, 137.0, 132.4, 130.2, 128.1, 124.6, 100.91, 100.9, 76.1, 73.3, 72.71, 72.7, 71.8, 70.9, 69.0, 68.61, 68.6, 66.6, 65.5, 62.1, 52.3, 38.9, 38.4, 35.9, 32.5, 32.1, 29.82, 29.81, 29.8, 29.6, 29.5, 29.3, 29.1, 28.5, 27.21, 27.2, 22.8, 21.8, 21.0, 20.9, 20.8, 20.7, 20.61, 20.6, 14.3. HR-ESI-MS m/z: [M + CH3OH–H]− Calcd for C67H108NO24S 1342.6982; Found: 1342.7032.
3.10. (2S,3R,E)-2-Octadecanamido-3-pivaloyloxy-octadec-4-en-1-yl 2,3,4-tri-O-acetyl-6-O-(p-toluenesulfonyl)-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-D-glucopyranoside (13a)
To a stirred solution of 12a (59 mg, 48.58 µmol) in dry DCM (10 mL) was added TFA (1.0 mL) dropwise over 5 min in an ice bath. After the ice bath was removed, the mixture was stirred at rt for 2.5 h. Toluene was added, and the organic layer was washed with NaHCO3 solution three times, dried over Na2SO4, and filtered. The filtrate was concentrated under reduced pressure, affording the free amine derivative. To the solution of stearic acid (20.2 mg, 70.9 µmol) dissolved in DCM (8.7 mL) in an ice-bath was added EDC (13.6 mg, 70.9 µmol) under an N2 atmosphere. After the mixture was stirred for 15 min, the above-obtained free amine (39.5 mg, 35.45 µmol) in DCM (4.3 mL) was added dropwise followed by the addition of DMAP (0.8 mg, 7.1 µmol). The reaction mixture was stirred at rt for 24 h. NaHCO3 solution was added dropwise to the mixture at 0 °C, and the aqueous layer was extracted with DCM three times. The combined organic layer was concentrated under vacuo, and the residue was purified using silica gel column chromatography to afford 13a (65%, 43.6 mg) as an off-white solid. TLC: Rf = 0.55 (EtOAc:Hex 50:50). 1H NMR (600 MHz, CDCl3): δ 7.78 (d, J = 8.3 Hz, 2H), 7.38 (d, J = 8.1 Hz, 2H), 5.75 (dt, J = 14.8, 7.4 Hz, 1H, =CH-), 5.61 (d, J = 9.3 Hz, 1H, −NHCO−), 5.36–5.33 (m, 2H, =CH- and 4′-H), 5.23 (t, J = 7.1 Hz, 1H), 5.19 (t, J = 9.3 Hz, 1H), 5.06 (dd, J = 10.3, 8.0 Hz, 1H), 4.92 (dd, J = 10.4, 3.4 Hz, 1H), 4.86 (dd, J = 9.4, 7.9 Hz, 1H), 4.47 (d, J = 7.9 Hz, 1H, anomeric), 4.45–4.44 (m, 1H), 4.43 (d, J = 7.8 Hz, 1H, anomeric), 4.33 (ddd, J = 11.3, 8.7, 4.1 Hz, 1H), 4.06 (ddd, J = 16.9, 11.1, 5.6 Hz, 1H), 3.99 (dd, J = 10.2, 6.5 Hz, 1H), 3.90 (dd, J = 12.6, 5.6 Hz, 1H), 3.78 (t, J = 9.5 Hz, 1H), 3.60 (ddd, J = 9.9, 4.7, 2.0 Hz, 1H), 3.52 (dd, J = 10.0, 4.4 Hz, 1H), 2.47 (s, 3H), 2.11 (s, 3H), 2.06 (s, 3H), 2.04 (s, 3H), 2.03 (s, 3H), 2.00 (s, 3H), 1.95 (s, 3H), 1.59–1.56 (m, 2H), 1.25 (s, 56H), 1.17 (s, 9H), 0.88 (t, J = 7.0 Hz, 6H, 2 × Me). 13C NMR (151 MHz, CDCl3): δ 177.1, 172.7, 170.5, 170.01, 170.0, 169.81, 169.8, 169.2, 145.7, 137.1, 132.4, 130.2, 128.2, 124.9, 100.9, 100.6, 76.0, 73.2, 72.8, 72.5, 72.0, 71.0, 70.9, 69.1, 67.8, 66.7, 65.6, 62.0, 50.6, 38.9, 37.0, 32.5, 32.1, 29.9, 29.8, 29.71, 29.7, 29.6, 29.5, 29.4, 29.2, 27.2, 25.9, 22.9, 21.8, 21.0, 20.91, 20.9, 20.8, 20.7, 20.6, 14.3. HR-ESI-MS m/z: [M + Na]+ Calcd for C72H117NNaO22S: 1402.7686; Found: 1402.7698.
3.11. (2S,3R,E)-2-Pentacosanamido-3-pivaloyloxy-octadec-4-en-1-yl 2,3,4-tri-O-acetyl-6-O-(p-toluenesulfonyl)-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-D-glucopyranoside (13b)
Compound 13b (64%, 44.4 mg) was prepared as a white solid from 12b (57 mg, 46.9 µmol) by the same method and conditions employed to prepare 13a. TLC: Rf = 0.5 (EtOAc:Hex 50:50). 1H NMR (600 MHz, CDCl3): δ 7.78 (d, J = 8.3 Hz, 2H), 7.38 (d, J = 8.1 Hz, 2H), 5.75 (dt, J = 14.7, 7.3 Hz, 1H, =CH-), 5.61 (d, J = 9.2 Hz, 1H, -NHCO-), 5.36–5.33 (m, 2H, =CH- and 4′-H), 5.23 (t, J = 7.1 Hz, 1H), 5.18 (t, J = 9.3 Hz, 1H), 5.06 (dd, J = 10.3, 8.0 Hz, 1H), 4.92 (dd, J = 10.4, 3.4 Hz, 1H), 4.86 (dd, J = 9.4, 7.9 Hz, 1H), 4.47 (d, J = 7.9 Hz, 1H, anomeric), 4.45 (d, J = 1.6 Hz, 1H), 4.43 (d, J = 7.8 Hz, 1H, anomeric), 4.33 (ddd, J = 11.3, 8.8, 4.1 Hz, 1H), 4.06 (ddd, J = 16.8, 11.1, 5.6 Hz, 2H), 3.99 (dd, J = 10.2, 6.6 Hz, 1H), 3.90 (dd, J = 12.0, 5.5 Hz, 2H), 3.78 (t, J = 9.5 Hz, 1H), 3.60 (ddd, J = 9.8, 4.7, 1.9 Hz, 1H), 3.52 (dd, J = 10.0, 4.4 Hz, 1H), 2.47 (s, 3H), 2.11 (s, 3H), 2.06 (s, 3H), 2.04 (s, 3H), 2.03 (s, 3H), 1.99 (s, 3H), 1.95 (s, 3H), 1.59–1.56 (m, 2H), 1.32–1.22 (m, 66H), 1.17 (s, 9H), 0.88 (t, J = 7.0 Hz, 6H, 2 × Me). 13C NMR (151 MHz, CDCl3): δ 177.1, 172.7, 170.5, 170.01, 170.0, 169.81, 169.8, 169.2, 145.6, 137.1, 132.4, 130.2, 128.2, 124.9, 100.9, 100.6, 76.0, 73.2, 72.8, 72.5, 72.0, 71.0, 70.9, 69.1, 67.8, 66.7, 65.6, 62.1, 50.6, 38.9, 37.0, 32.5, 32.1, 29.91, 29.9, 29.8, 29.71, 29.7, 29.6, 29.51, 29.5, 29.4, 29.2, 27.2, 25.9, 22.9, 21.8, 21.0, 20.91, 20.9, 20.8, 20.7, 20.6, 14.3. HR-ESI-MS m/z: [M + HCOO]− Calcd for C80H132NO24S 1522.8860; found: 1522.8896.
3.12. (2S,3R,E)-2-Octadecanamido-3-pivaloyloxy-pentacos-4-en-1-yl 2,3,4-tri-O-acetyl-6-O-(p-toluenesulfonyl)-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-D-glucopyranoside (13c)
Compound 13c (63%, 36.2 mg) as a white solid was prepared from 12c (51 mg, 38.9 µmol) by the same method and conditions employed to prepare 13a. TLC: Rf = 0.5 (EtOAc:Hex 50:50). 1H NMR (600 MHz, CDCl3): δ 7.78 (d, J = 8.3 Hz, 2H), 7.38 (d, J = 8.0 Hz, 2H), 5.75 (dt, J = 14.7, 7.3 Hz, 1H, =CH-), 5.61 (d, J = 9.2 Hz, 1H, -NHCO-), 5.36–5.33 (m, 2H, =CH- and 4′-H), 5.23 (t, J = 7.2 Hz, 1H), 5.18 (t, J = 9.3 Hz, 1H), 5.06 (dd, J = 10.4, 7.9 Hz, 1H), 4.92 (dd, J = 10.4, 3.4 Hz, 1H), 4.86 (dd, J = 9.4, 7.9 Hz, 1H), 4.47 (d, J = 7.9 Hz, 1H, anomeric), 4.45 (d, J = 1.6 Hz, 1H), 4.43 (d, J = 7.8 Hz, 1H, anomeric), 4.33 (ddd, J = 11.3, 8.8, 4.2 Hz, 1H), 4.06 (ddd, J = 16.9, 11.1, 5.6 Hz, 2H), 3.99 (dd, J = 10.2, 6.6 Hz, 1H), 3.93–3.88 (m, 2H), 3.78 (t, J = 9.5 Hz, 1H), 3.60 (ddd, J = 9.9, 4.7, 2.0 Hz, 1H), 3.52 (dd, J = 10.0, 4.4 Hz, 1H), 2.47 (s, 3H), 2.11 (s, 3H), 2.06 (s, 3H), 2.04 (s, 3H), 2.03 (s, 3H), 1.99 (s, 3H), 1.95 (s, 3H), 1.57 (b, 2H), 1.35–1.22 (m, 70H), 1.17 (s, 9H), 0.88 (t, J = 7.0 Hz, 6H, 2 × Me). 13C NMR (151 MHz, CDCl3): δ 177.1, 172.7, 170.5, 170.01, 170.0, 169.81, 169.8, 169.2, 145.6, 137.2, 132.4, 130.2, 128.2, 124.9, 100.9, 100.6, 76.0, 73.1, 72.8, 72.5, 72.0, 71.0, 70.9, 69.1, 67.8, 66.7, 65.6, 62.0, 50.6, 38.9, 37.0, 32.5, 32.1, 29.9, 29.8, 29.71, 29.7, 29.6, 29.5, 29.4, 29.2, 27.2, 25.9, 22.9, 21.8, 21.0, 20.91, 20.9, 20.8, 20.7, 20.6, 14.3. HR-ESI-MS m/z: [M + CH3OH-H]− Calcd for C80H134NO23S 1508.9067; found: 1508.9098.
3.13. (2S,3R,E)-2-Pentacosanamido-3-pivaloyloxy-pentacos-4-en-1-yl 2,3,4-tri-O-acetyl-6-O-(p-toluenesulfonyl)-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-D-glucopyranoside (13d)
Compound 13d (63%, 37.8 mg) was prepared as a white solid from 12d (50 mg, 38.1 µmol) by the same method and conditions employed to prepare 13a. TLC: Rf = 0.5 (EtOAc:Hex 50:50). 1H NMR (600 MHz, CDCl3): δ 7.77 (d, J = 8.3 Hz, 2H), 7.38 (d, J = 8.1 Hz, 2H), 5.75 (dt, J = 14.8, 7.3 Hz, 1H, =CH-), 5.61 (d, J = 9.2 Hz, 1H, -NHCO-), 5.36–5.33 (m, 2H, =CH- and 4′-H), 5.23 (t, J = 7.1 Hz, 1H), 5.18 (t, J = 9.3 Hz, 1H), 5.06 (dd, J = 10.3, 8.0 Hz, 1H), 4.92 (dd, J = 10.4, 3.4 Hz, 1H), 4.86 (dd, J = 9.4, 7.9 Hz, 1H), 4.47 (d, J = 7.9 Hz, 1H, anomeric), 4.45 (d, J = 1.4 Hz, 1H), 4.43 (d, J = 7.8 Hz, 1H, anomeric), 4.33 (ddd, J = 11.3, 8.7, 4.1 Hz, 1H), 4.05 (ddd, J = 16.9, 11.1, 5.6 Hz, 2H), 3.99 (dd, J = 10.2, 6.5 Hz, 1H), 3.90 (dd, J = 11.9, 5.5 Hz, 2H), 3.78 (t, J = 9.5 Hz, 1H), 3.60 (ddd, J = 9.9, 4.7, 2.0 Hz, 1H), 3.52 (dd, J = 10.0, 4.4 Hz, 1H), 2.47 (s, 3H), 2.11 (s, 3H), 2.06 (s, 3H), 2.04 (s, 3H), 2.03 (s, 3H), 1.99 (s, 3H), 1.95 (s, 3H), 1.64 (dd, J = 14.7, 7.2 Hz, 2H), 1.59–1.56 (m, 2H), 1.34–1.23 (m, 80H), 1.17 (s, 9H), 0.88 (t, J = 7.0 Hz, 6H, 2 × Me). 13C NMR (151 MHz, CDCl3): δ 177.1, 172.7, 170.5, 170.01, 170.0, 169.81, 169.8, 169.2, 145.6, 137.2, 132.4, 130.2, 128.2, 124.9, 100.9, 100.6, 76.0, 73.1, 72.8, 72.5, 72.0, 71.0, 70.9, 69.1, 67.8, 66.7, 65.6, 62.0, 50.6, 38.9, 37.0, 32.5, 32.1, 29.91, 29.9, 29.8, 29.71, 29.7, 29.6, 29.5, 29.4, 29.2, 27.2, 25.9, 22.9, 21.8, 21.0, 20.91, 20.9, 20.8, 20.7, 20.6, 14.3. HR-ESI-MS m/z: [M-H]− Calcd for C86H144NO22S 1574.9901; found: 1574.9834.
3.14. (2S,3R,E)-2-Octadecanamido-3-pivaloyloxy-octadec-4-en-1-yl 2,3,4-tri-O-acetyl-6-azido-6-deoxy-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-D-glucopyranoside (14a)
To a solution of 13a (30 mg, 21.7 µmol) in dry DMF (6.0 mL) was added NaN3 (28 mg, 434.5 µmol), and the mixture was heated at 86 °C for 2 d. When the starting material 13a disappeared in TLC, the reaction was allowed to cool to rt. The mixture was diluted with EtOAc and cooled at 0 °C. After cold water was added, the mixture was extracted with EtOAc three times. The combined organic layers were washed with brine, filtered, dried over Na2SO4, and condensed under vacuum. The residue was purified through silica gel column chromatography to afford 14a as a white solid (80%, 21.8 mg). TLC: Rf = 0.45 (EtOAc:Hex 40:60). 1H NMR (600 MHz, CDCl3): δ 5.75 (dt, J = 14.8, 7.4 Hz, 1H, =CH-), 5.62 (d, J = 9.2 Hz, 1H, −NHCO−), 5.38–5.30 (m, 2H, =CH- and 4′-H), 5.25–5.17 (m, 2H), 5.09 (dd, J = 10.4, 7.9 Hz, 1H), 4.96 (dd, J = 10.4, 3.5 Hz, 1H), 4.88 (dd, J = 9.5, 7.8 Hz, 1H), 4.49–4.44 (m, 1H), 4.50 (d, J = 7.9 Hz, 1H, anomeric), 4.42 (d, J = 7.7 Hz, 1H, anomeric), 4.32 (ddd, J = 11.4, 8.6, 4.1 Hz, 1H), 4.05 (dd, J = 12.0, 4.9 Hz, 1H), 3.90 (dd, J = 10.0, 3.7 Hz, 1H), 3.85 (t, J = 9.5 Hz, 1H), 3.72 (t, J = 6.7 Hz, 1H), 3.60 (ddd, J = 9.9, 4.7, 2.0 Hz, 1H), 3.52 (dd, J = 10.0, 4.4 Hz, 1H), 3.47 (dd, J = 12.8, 7.3 Hz, 1H), 3.26 (dd, J = 12.8, 5.6 Hz, 1H), 2.17 (s, 3H), 2.12 (s, 3H), 2.07 (s, 3H), 2.04 (s, 3H), 2.04 (s, 3H), 1.97 (s, 3H), 1.59–1.56 (b, 2H), 1.35–1.22 (m, 56H), 1.17 (s, 9H), 0.88 (t, J = 7.0 Hz, 6H, 2 × Me). 13C NMR (151 MHz, CDCl3): δ 177.1, 172.7, 170.5, 170.2, 170.1, 169.81, 169.8, 169.2, 137.2, 125.0, 100.8, 100.7, 75.4, 73.2, 72.9, 72.5, 72.4, 72.0, 71.1, 69.3, 69.2, 67.8, 67.6, 62.0, 50.6, 50.4, 38.9, 37.0, 32.5, 32.1, 29.9, 29.8, 29.71, 29.7, 29.6, 29.5, 29.4, 29.2, 27.2, 25.9, 22.9, 21.1, 21.0, 20.9, 20.8, 20.7, 14.3. HR-ESI-MS m/z: [M + HCOO]− Calcd for C66H111N4O21: 1295.7741; Found: 1295.7753.
3.15. (2S,3R,E)-2-Pentacosanamido-3-pivaloyloxy-octadec-4-en-1-yl 2,3,4-tri-O-acetyl-6-azido-6-deoxy-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-D-glucopyranoside (14b)
Compound 14b (76%, 20.8 mg) was prepared as a white solid from 13b (30 mg, 20.3 µmol) by the same method and conditions employed to prepare 14a. TLC: Rf = 0.65 (EtOAc:Hex 40:60). 1H NMR (600 MHz, CDCl3): δ 5.75 (dt, J = 14.7, 7.4 Hz, 1H, =CH-), 5.61 (d, J = 9.2 Hz, 1H, -NHCO-), 5.37–5.31 (m, 2H, =CH- and 4′-H), 5.21 (dt, J = 15.5, 8.3 Hz, 2H), 5.09 (dd, J = 10.4, 7.9 Hz, 1H), 4.96 (dd, J = 10.4, 3.5 Hz, 1H), 4.88 (dd, J = 9.5, 7.8 Hz, 1H), 4.51 (d, J = 7.8 Hz, 1H, anomeric), 4.48 (d, J = 1.7 Hz, 1H), 4.42 (d, J = 7.7 Hz, 1H, anomeric), 4.32 (ddd, J = 11.4, 8.4, 4.1 Hz, 1H), 4.05 (dd, J = 12.0, 4.9 Hz, 1H), 3.91 (dd, J = 9.9, 3.7 Hz, 1H), 3.85 (t, J = 9.5 Hz, 1H), 3.72 (t, J = 6.6 Hz, 1H), 3.60 (ddd, J = 9.9, 4.7, 2.0 Hz, 1H), 3.52 (dd, J = 10.0, 4.4 Hz, 1H), 3.47 (dd, J = 12.8, 7.3 Hz, 1H), 3.26 (dd, J = 12.8, 5.6 Hz, 1H), 2.17 (s, 3H), 2.12 (s, 3H), 2.07 (s, 3H), 2.04 (s, 3H), 2.04 (s, 3H), 1.97 (s, 3H), 1.59–1.56 (b, 2H), 1.37–1.21 (m, 70H), 1.17 (s, 9H), 0.88 (t, J = 7.0 Hz, 6H, 2 × Me). 13C NMR (151 MHz, CDCl3): δ 177.1, 172.7, 170.4, 170.2, 170.1, 169.81, 169.8, 169.2, 137.2, 125.0, 100.8, 100.7, 75.4, 73.2, 72.9, 72.5, 72.4, 72.0, 71.1, 69.2, 67.8, 67.6, 62.4, 62.2, 50.6, 50.4, 38.9, 37.0, 32.5, 32.1, 29.9, 29.8, 29.71, 29.7, 29.6, 29.5, 29.4, 29.2, 27.2, 25.9, 22.9, 21.1, 21.0, 20.9, 20.8, 20.7, 14.3. HR-ESI-MS m/z: [M + CH3OH-H]- Calcd for C73H127N4O20 1379.9044; Found: 1379.9058.
3.16. (2S,3R,E)-2-Octadecanamido-3-pivaloyloxy-pentacos-4-en-1-yl 2,3,4-tri-O-acetyl-6-azido-6-deoxy-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-D-glucopyranoside (14c)
Compound 14c (69%, 18.9 mg) was prepared as a white solid from 13c (30 mg, 20.3 µmol) by the same method and conditions employed to prepare 14a. TLC: Rf = 0.45 (EtOAc:Hex 40:60). 1H NMR (600 MHz, CDCl3) δ 5.75 (dt, J = 14.7, 7.4 Hz, 1H, =CH-), 5.62 (d, J = 9.3 Hz, 1H, -NHCO-), 5.35–5.32 (m, 2H, =CH- and 4′-H), 5.24–5.16 (m, 2H), 5.09 (dd, J = 10.4, 7.9 Hz, 1H), 4.96 (dd, J = 10.4, 3.5 Hz, 1H), 4.87 (dd, J = 9.5, 7.8 Hz, 1H), 4.50 (d, J = 7.8 Hz, 1H, anomeric), 4.48 (d, J = 1.8 Hz, 1H), 4.42 (d, J = 7.7 Hz, 1H, anomeric), 4.32 (ddd, J = 11.4, 8.4, 4.1 Hz, 1H), 4.05 (dd, J = 12.0, 4.9 Hz, 1H), 3.90 (dd, J = 10.0, 3.7 Hz, 1H), 3.85 (t, J = 9.5 Hz, 1H), 3.72 (t, J = 6.6 Hz, 1H), 3.60 (ddd, J = 9.8, 4.7, 2.0 Hz, 1H), 3.51 (dd, J = 10.0, 4.4 Hz, 1H), 3.46 (dd, J = 12.8, 7.3 Hz, 1H), 3.26 (dd, J = 12.8, 5.6 Hz, 1H), 2.16 (s, 3H), 2.11 (s, 3H), 2.07 (s, 3H), 2.04 (s, 6H), 1.97 (s, 3H), 1.59–1.56 (b, 2H), 1.34–1.23 (m, 70H), 1.17 (s, 9H), 0.88 (t, J = 7.0 Hz, 6H, 2 × Me). 13C NMR (151 MHz, CDCl3): δ 177.1, 172.7, 170.4, 170.2, 170.1, 169.81, 169.8, 169.2, 137.2, 125.0, 100.71, 100.7, 75.4, 73.2, 72.9, 72.5, 72.4, 71.9, 71.1, 69.2, 67.8, 67.6, 62.1, 62.0, 50.6, 50.4, 38.9, 37.0, 32.5, 32.1, 29.9, 29.81, 29.8, 29.71, 29.7, 29.6, 29.5, 29.4, 29.2, 27.2, 25.9, 22.9, 21.1, 21.0, 20.9, 20.8, 20.7, 14.3. HR-ESI-MS m/z: [M + HCOO]− Calcd for C73H125N4O21 1393.8837; Found: 1393.8878.
3.17. (2S,3R,E)-2-Pentacosanamido-3-pivaloyloxy-pentacos-4-en-1-yl 2,3,4-tri-O-acetyl-6-azido-6-deoxy-β-D-galactopyranosyl-(1→4)-2,3,6-tri-O-acetyl-D-glucopyranoside (14d)
Compound 14d (72%, 18.5 mg) was prepared as a white solid from 13d (28 mg, 17.8 µmol) by the same method and conditions employed to prepare 14a. TLC: Rf = 0.66 (EtOAc:Hex 40:60). 1H NMR (600 MHz, CDCl3): δ 5.75 (dt, J = 14.7, 7.4 Hz, 1H, =CH-), 5.61 (d, J = 9.2 Hz, 1H, -NHCO-), 5.36–5.32 (m, 2H, =CH- and 4′-H), 5.24–5.17 (m, 2H), 5.09 (dd, J = 10.4, 7.9 Hz, 1H), 4.96 (dd, J = 10.4, 3.5 Hz, 1H), 4.87 (dd, J = 9.5, 7.8 Hz, 1H), 4.50 (d, J = 7.9 Hz, 1H, anomeric), 4.48 (d, J = 1.7 Hz, 1H), 4.42 (d, J = 7.7 Hz, 1H, anomeric), 4.32 (ddd, J = 11.4, 8.2, 4.1 Hz, 1H), 4.05 (dd, J = 12.0, 4.9 Hz, 1H), 3.90 (dd, J = 9.9, 3.7 Hz, 1H), 3.85 (t, J = 9.5 Hz, 1H), 3.72 (t, J = 6.5 Hz, 1H), 3.60 (ddd, J = 9.8, 4.7, 2.0 Hz, 1H), 3.51 (dd, J = 10.0, 4.4 Hz, 1H), 3.47 (dd, J = 12.8, 7.3 Hz, 1H), 3.26 (dd, J = 12.8, 5.7 Hz, 1H), 2.17 (s, 3H), 2.12 (s, 3H), 2.07 (s, 3H), 2.04 (s, 6H, 2 × CH3), 1.97 (s, 3H), 1.59–1.56 (b, 2H), 1.32–1.23 (m, 84H), 1.17 (s, 9H), 0.88 (t, J = 7.0 Hz, 6H, 2 × Me). 13C NMR (151 MHz, CDCl3): δ 177.1, 172.7, 170.4, 170.2, 170.1, 169.81, 169.8, 169.2, 137.2, 125.0, 100.8, 100.7, 75.4, 73.2, 72.9, 72.5, 72.4, 72.0, 71.1, 69.2, 67.8, 67.6, 63.2, 62.0, 50.6, 50.4, 38.9, 37.0, 32.5, 32.1, 29.9, 29.89, 29.8, 29.71, 29.7, 29.6, 29.5, 29.4, 29.2, 27.2, 25.9, 22.9, 21.1, 20.91, 20.9, 20.8, 20.7, 14.3. HR-ESI-MS m/z: [M + CH3OH-H]− Calcd for C80H141N4O20 1478.0139; Found: 1478.0077.
3.18. (2S,3R,E)-2-Octadecanamido-3-hydroxy-octadec-4-en-1-yl 6-azido-6-deoxy-β-D-galactopyranosyl-(1→4)-β-D-glucopyranoside (1a)
To a solution of 14a (19.5 mg, 15.6 µmol) in a mixture of dry DCM and MeOH (1:1, 2 mL) was added NaOMe in MeOH (5 M, 43.6 µL, 0.2 mmol) dropwise at 0 °C. The ice bath was removed, and the mixture was stirred at rt under N2 for 2 d. After the starting material was completely consumed as indicated by TLC, Amberlyst H+ resin was added with stirring to neutralize the reaction mixture. The solvent was removed under vacuum, and the residue was subjected to silica gel column chromatography to afford 1a as an off-white solid (90%, 12.8 mg). TLC: Rf = 0.51 (DCM:MeOH 85:15). 1H NMR (600 MHz, CDCl3:CD3OD 2:1): δ 5.65 (dt, J = 15.0, 7.4 Hz, 1H, =CH-), 5.41 (dd, J = 15.0, 7.6 Hz, 1H, =CH-), 4.32 (d, J = 7.8 Hz, 1H anomeric), 4.25 (d, J = 7.8 Hz, 1H anomeric), 4.17 (dd, J = 10.0, 3.9 Hz, 1H), 4.05 (t, J = 7.6 Hz, 1H), 3.96–3.93 (m, 1H), 3.82 (d, J = 3.0 Hz, 2H), 3.77 (d, J = 2.9 Hz, 1H), 3.65–3.61 (m, 2H), 3.60–3.58 (m, 1H), 3.56–3.52 (m, 4H), 3.50 (dd, J = 9.8, 3.0 Hz, 2H), 3.47 (dd, J = 9.7, 3.2 Hz, 1H), 3.38–3.35 (m, 1H), 2.13 (t, J = 7.7 Hz, 2H), 2.00–1.96 (m, 2H), 1.56–1.52 (m, 2H), 1.34–1.30 (m, 2H), 1.28–1.20 (m, 48H), 0.84 (t, J = 7.0 Hz, 6H, 2 × Me). 13C NMR (151 MHz, CDCl3:CD3OD 2:1): δ 175.0, 134.8, 129.7, 104.3, 103.3, 80.4, 75.3, 75.2, 74.0, 73.8, 73.7, 72.4, 71.3, 69.2, 69.0, 61.2, 53.6, 51.6, 36.9, 32.8, 32.3, 30.12, 30.11, 30.1, 30.03, 30.02, 30.01, 30.0, 29.9, 29.8, 29.72, 29.71, 29.7, 26.4, 23.0, 14.3. HR-ESI-MS m/z: [M + HCOO]− Calcd for C49H91N4O14 959.6532; Found: 959.6563.
3.19. (2S,3R,E)-2-Pentacosanamido-3-(hydroxy)-octadec-4-en-1-yl 6-azido-6-deoxy-β-D-galactopyranosyl-(1→4)-β-D-glucopyranoside (1b)
Compound 1b (88%, 11.3 mg) was prepared as a white solid from 14b (17 mg, 12.6 µmol) by the same method and conditions utilized to prepare 1a. TLC: Rf = 0.66 (DCM:MeOH 85:15). 1H NMR (600 MHz, CDCl3:CD3OD 2:1): δ 5.63 (dt, J = 14.9, 7.2 Hz, 1H, =CH-), 5.40 (dd, J = 14.9, 7.5 Hz, 1H, =CH-), 4.30 (d, J = 7.8 Hz, 1H anomeric), 4.23 (d, J = 7.8 Hz, 1H anomeric), 4.14 (dd, J = 10.1, 3.9 Hz, 1H), 4.04 (t, J = 7.4 Hz, 1H), 3.80 (d, J = 3.0 Hz, 2H), 3.76 (d, J = 3.0 Hz, 1H), 3.61–3.59 (m, 2H), 3.58–3.56 (m, 4H), 3.54–3.51 (m, 4H), 3.49 (dd, J = 10.4, 1.9 Hz, 1H), 3.46 (dd, J = 9.7, 3.2 Hz, 1H), 2.11 (t, J = 7.7 Hz, 2H), 1.98–1.94 (m, 2H), 1.54–1.50 (m, 2H), 1.31–1.28 (m, 2H), 1.26–1.19 (m, 62H), 0.83 (t, J = 7.0 Hz, 6H, 2 × Me). 13C NMR (151 MHz, CDCl3:CD3OD 2:1): δ 174.6, 134.6, 129.2, 104.0, 103.0, 80.2, 75.0, 74.9, 73.7, 73.4, 72.3, 71.0, 68.8, 68.7, 63.6, 61.0, 53.5, 53.3, 51.3, 36.6, 32.5, 32.0, 29.82, 29.81, 29.8, 29.71, 29.7, 29.6, 29.52, 29.51, 29.5, 29.4, 26.0, 22.8, 14.1. HR-ESI-MS m/z: [M + HCOO]− Calcd for C56H105N4O14 1057.7628; Found: 1057.7659.
3.20. (2S,3R,E)-2-Octadecanamido-3-(hydroxy)-pentacos-4-en-1-yl 6-azido-6-deoxy-β-D-galactopyranosyl-(1→4)-β-D-glucopyranoside (1c)
Compound 1c (80%, 10.8 mg) was prepared as an off-white solid from 14c (18 mg, 13.3 µmol) by the same method and conditions utilized to prepare 1a. TLC: Rf = 0.70 (DCM:MeOH 85:15). 1H NMR (600 MHz, CDCl3:CD3OD 2:1): δ 5.64 (dt, J = 14.9, 7.3 Hz, 1H, =CH-), 5.41 (dd, J = 14.9, 7.6 Hz, 1H, =CH-), 4.30 (1H, anomeric, overlapped with DHO signal), 4.25 (d, J = 7.8 Hz, 1H, anomeric), 4.16 (dd, J = 10.0, 3.9 Hz, 1H), 4.05 (t, J = 7.7 Hz, 1H), 3.96–3.92 (m, 2H), 3.92–3.89 (m, 3H), 3.82 (d, J = 3.3 Hz, 2H), 3.77 (d, J = 3.3 Hz, 1H), 3.63–3.58 (m, 2H), 3.55–3.52 (m, 3H), 3.50 (dd, J = 10.5, 3.3 Hz, 1H), 3.47 (dd, J = 9.7, 3.2 Hz, 1H), 2.15–2.10 (m, 2H), 1.99–1.96 (m, 2H), 1.56–1.52 (m, 2H), 1.34–1.29 (m, 2H), 1.27–1.20 (m, 62H), 0.84 (t, J = 7.0 Hz, 6H, 2 × Me). 13C NMR (151 MHz, CDCl3:CD3OD 2:1): δ 174.9, 134.8, 129.6, 104.2, 103.2, 80.4, 75.2, 75.1, 74.0, 73.7, 73.6, 72.4, 71.2, 70.6, 69.1, 61.1, 55.3, 53.5, 51.5, 49.9, 49.6, 36.8, 32.7, 32.2, 30.1, 30.03, 30.02, 30.01, 30.0, 29.91, 29.9, 29.8, 29.73, 29.72, 29.71, 29.7, 29.6, 26.3, 23.0, 14.2. HR-ESI-MS m/z: [M + HCOO]− Calcd for C56H105N4O14 1057.7628; Found: 1057.7659.
3.21. (2S,3R,E)-2-Pentacosanamido-3-(hydroxy)-pentacos-4-en-1-yl 6-azido-6-deoxy-β-D-galactopyranosyl-(1→4)-β-D-glucopyranoside (1d)
Compound 1d (85%, 11.1 mg) was prepared as a white solid from 14d (17 mg, 11.7 µmol) by the same method and conditions used to prepare 1a. TLC: Rf = 0.75 (DCM:MeOH 85:15). 1H NMR (600 MHz, CDCl3:CD3OD 2:1): δ 5.63 (dt, J = 14.9, 7.3 Hz, 1H, =CH-), 5.39 (dd, J = 14.9, 7.5 Hz, 1H, =CH-), 4.30 (d, J = 7.8 Hz, 1H, anomeric), 4.23 (d, J = 7.8 Hz, 1H, anomeric), 4.13 (dd, J = 10.0, 4.0 Hz, 1H), 4.04 (t, J = 7.4 Hz, 1H), 3.94–3.90 (m, 1H), 3.79 (d, J = 2.7 Hz, 3H), 3.61–3.56 (m, 3H), 3.54–3.52 (m, 2H), 3.51 (dd, J = 7.8, 3.1 Hz, 2H), 3.48 (dd, J = 10.7, 3.2 Hz, 1H), 3.44–3.44 (m, 3H), 2.11 (t, J = 7.6 Hz, 2H), 1.94–1.97 (m, 2H), 1.59–1.47 (m, 4H), 1.31–1.16 (m, 76H), 0.82 (t, J = 7.0 Hz, 6H). 13C NMR (151 MHz, CDCl3:CD3OD 2:1): δ 177.6, 134.5, 129.0, 103.8, 102.8, 79.9, 74.8, 73.6, 73.21, 73.2, 72.2, 70.8, 68.61, 68.6, 63.5, 60.9, 53.1, 51.1, 36.5, 32.3, 31.9, 29.71, 29.7, 29.61, 29.6, 29.51, 29.5, 29.42, 29.41, 29.4, 29.31, 29.3, 29.21, 29.2, 25.9, 22.6, 14.0. HR-ESI-MS m/z: [M + HCOO]− Calcd for C63H119N4O14 1155.8723; Found: 1155.87622.1.
4. Conclusions
β-LacCer is not only a key intermediate in complex GSL biosynthesis but also an important player in many biological events and diseases. However, in-depth investigation of β-LacCer is hindered by its lack of fluorophores or other useful functionalities. To overcome this problem, we designed and synthesized a series of β-LacCer analogs carrying an azido group in its glycan and varied Cer moieties. In the literature, there are many reports [33,34,35,36,37,38,39,40,41,42,43,44,45,46] (just cite a few) about the synthesis of β-LacCer and its analogs, but none of these reports have described azide-functionalized β-LacCer analogs like 1a–d in this work. To facilitate rapid access to 1a–d, we have employed a diversity-oriented synthetic strategy that is highlighted by the late-stage lipid remodeling for on-site assembly of the Cer moiety. Therefore, all the target molecules 1a–d could be readily derived from the same intermediate 10 within five robust steps. This strategy and its protocols are streamlined so that they will be applicable to the synthesis of various lipid forms of β-LacCer and its analogs. The azide in 1a–d is suitable for further and direct functionalization by means of straightforward and biocompatible click chemistry to attach a diversity of molecular tags, such as affinity and fluorescent tags that enable different modern analytical technologies. Alternatively, the azido group in 1a–d can be reduced to generate a free primary amino group that is suitable for further introduction of molecular labels via chemoselective N-acylation. Moreover, additional functionalization of 1a–d through biocompatible click chemistry can be conducted either before or after their introduction to cells/tissues [24,25], thereby expanding their application scopes. The flexibility to introduce fluorophores or other large molecular labels later also makes 1a–d especially valuable. Because having a hydroxyl group substituted with a small azido group will have a minimal impact on the interaction of these probes with target molecules in/on cells and probably make the probes more easily acceptable by enzymes, these probes can better mimic the behaviors and bioactivities of β-LacCer than those containing large functional groups. Consequently, this work not only has established a novel and efficient method for the synthesis of β-LacCer and its analogs but also has provided 1a–d as powerful probes or tools for in-depth investigation of β-LacCer biology, including the influences of its lipid structures on the biological functions and metabolisms of β-LacCer and other GSLs, as well as their related diseases.
Supplementary Materials
The following supporting information can be downloaded at: https://www.mdpi.com/article/10.3390/molecules30132667/s1, the NMR and MS spectra of all new compounds, including 4, 5, 6, 7, 8 (only NMR), 10, 12a,b, 13a–d, 14a–d, and 1a–d.
Author Contributions
B.M. performed the experiments and data interpretation and drafted the manuscript; R.R. and X.Y. were engaged in experimental design and revision, provided advice, and verified data; A.M.G., N.A.O., and H.A.A.-F. provided research supervision and advice; Z.G. is overall responsible for this project, including experimental designs, data analysis, and research supervision. All authors have read and agreed to the published version of the manuscript.
Funding
We acknowledge the Egyptian Government, represented by the Egyptian Cultural and Educational Bureau at Washington DC, and the Mission Sector of Egyptian Ministry of Higher Education for supporting B.M. This work is partly supported by an NIH grant (R21 AI170129) to Z.G.
Data Availability Statement
The original contributions presented in this study are included in the article/Supplementary Material. Further inquiries can be directed to the corresponding author.
Acknowledgments
Z.G. is grateful to Steven and Rebecca Scott for their endowment to support our research.
Conflicts of Interest
The authors declare no conflict of interest.
References
- Guo, Z. The structural diversity of natural glycosphingolipids (GSLs). J. Carbohydr. Chem. 2022, 41, 63–154. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Hakomori, S.I. Structure and function of glycosphingolipids and sphingolipids: Recollections and future trends. Biochim. Biophys. Acta 2008, 1780, 325–346. [Google Scholar] [CrossRef] [Scilit]
- Ando, H.; Komura, N. Recent progress in the synthesis of glycosphingolipids. Curr. Opin. Chem. Biol. 2024, 78, 102423. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Sonnino, S.; Prinetti, A. Gangliosides as regulators of cell membrane organization and functions. Adv. Exp. Med. Biol. 2010, 688, 165–184. [Google Scholar] [PubMed]
- Saxena, K.; Zimmermann, P.; Schmidt, R.R.; Shipley, G.G. Bilayer properties of totally synthetic C16:0-lactosyl-ceramide. Biophys. J. 2000, 78, 306–312. [Google Scholar] [CrossRef] [Scilit]
- Sarmento, M.J.; Ricardo, J.C.; Amaro, M.; Šachl, R. Organization of gangliosides into membrane nanodomains. FEBS Lett. 2020, 594, 3668–3697. [Google Scholar] [CrossRef] [Scilit]
- Nakayama, H.; Ogawa, H.; Takamori, K.; Iwabuchi, K. GSL-enriched membrane microdomains in innate immune responses. Arch. Immunol. Ther. Exp. 2013, 61, 217–228. [Google Scholar] [CrossRef] [Scilit]
- Wennekes, T.; van den Berg, R.J.; Boot, R.G.; van der Marel, G.A.; Overkleeft, H.S.; Aerts, J.M. Glycosphingolipids—Nature, function, and pharmacological modulation. Angew. Chem. Int. Ed. Engl. 2009, 48, 8848–8869. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Hakomori, S. Cancer-associated glycosphingolipid antigens: Their structure, organization, and function. Acta Anat. 1998, 161, 79–90. [Google Scholar] [CrossRef] [Scilit]
- Mutoh, T.; Hirabayashi, Y.; Mihara, T.; Ueda, M.; Koga, H.; Ueda, A.; Kokura, T.; Yamamoto, H. Role of glycosphingolipids and therapeutic perspectives on Alzheimer’s disease. CNS Neurol. Disord. Drug Targets 2006, 5, 375–380. [Google Scholar] [CrossRef] [Scilit]
- Hakomori, S.; Zhang, Y. Glycosphingolipid antigens and cancer therapy. Chem. Biol. 1997, 3, 97–104. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Sandhoff, K.; Kolter, T. Biosynthesis and degradation of mammalian glycosphingolipids. Philos. Trans. R. Soc. Lond. B Biol. Sci. 2003, 358, 847–861. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Yu, R.K.; Tsai, Y.T.; Ariga, T.; Yanagisawa, M. Structures, biosynthesis, and functions of gangliosides—An overview. J. Oleo. Sci. 2011, 60, 537–544. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Yu, W.; Ying, J.; Wang, X.; Liu, X.; Zhao, T.; Yoon, S.; Zheng, Q.; Fang, Y.; Yang, D.; Hua, F. The Involvement of Lactosylceramide in Central Nervous System Inflammation Related to Neurodegenerative Disease. Front. Aging Neurosci. 2021, 13, 691230. [Google Scholar] [CrossRef] [Scilit]
- Nakayama, H.; Nagafuku, M.; Suzuki, A.; Iwabuchi, K.; Inokuchi, J.I. The regulatory roles of glycosphingolipid-enriched lipid rafts in immune systems. FEBS Lett. 2018, 592, 3921–3942. [Google Scholar] [CrossRef] [Scilit]
- Furukawa, K.; Ohmi, Y.; Kondo, Y.; Ohkawa, Y.; Tajima, O.; Furukawa, K. Regulatory function of glycosphingolipids in the inflammation and degeneration. Arch. Biochem. Biophys. 2015, 571, 58–65. [Google Scholar] [CrossRef] [Scilit]
- Silsirivanit, A.; Phoomak, C.; Teeravirote, K.; Wattanavises, S.; Seubwai, W.; Saengboonmee, C.; Zhan, Z.; Inokuchi, J.I.; Suzuki, A.; Wongkham, S. Overexpression of HexCer and LacCer containing 2-hydroxylated fatty acids in cholangiocarcinoma and the association of the increase of LacCer (d18:1-h23:0) with shorter survival of the patients. Glycoconj. J. 2019, 36, 103–111. [Google Scholar] [CrossRef] [Scilit]
- Bodas, M.; Min, T.; Vij, N. Lactosylceramide-accumulation in lipid-rafts mediate aberrant-autophagy, inflammation and apoptosis in cigarette smoke induced emphysema. Apoptosis 2015, 20, 725–739. [Google Scholar] [CrossRef] [Scilit]
- Chatterjee, S. Sphingolipids in atherosclerosis and vascular biology. Arter. Thromb. Vasc. Biol. 1998, 18, 1523–1533. [Google Scholar] [CrossRef] [Scilit]
- Chatterjee, S.; Kolmakova, A.; Rajesh, M. Regulation of lactosylceramide synthase (glucosylceramide beta1-->4 galactosyltransferase); implication as a drug target. Curr. Drug Targets 2008, 9, 272–281. [Google Scholar] [CrossRef] [Scilit]
- Ekyalongo, R.C.; Nakayama, H.; Kina, K.; Kaga, N.; Iwabuchi, K. Organization and functions of glycolipid-enriched microdomains in phagocytes. Biochim. Biophys. Acta 2015, 1851, 90–97. [Google Scholar] [CrossRef] [Scilit]
- Yoshizaki, F.; Nakayama, H.; Iwahara, C.; Takamori, K.; Ogawa, H.; Iwabuchi, K. Role of glycosphingolipid-enriched microdomains in innate immunity: Microdomain-dependent phagocytic cell functions. Biochim. Biophys. Acta 2008, 1780, 383–392. [Google Scholar] [CrossRef] [Scilit]
- Chiricozzi, E.; Ciampa, M.G.; Brasile, G.; Compostella, F.; Prinetti, A.; Nakayama, H.; Ekyalongo, R.C.; Iwabuchi, K.; Sonnino, S.; Mauri, L. Direct interaction, instrumental for signaling processes, between LacCer and Lyn in the lipid rafts of neutrophil-like cells. J. Lipid Res. 2015, 56, 129–141. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Kolb, H.C.; Finn, M.G.; Sharpless, K.B. Click chemistry: Diverse chemical function from a few good reactions. Angew. Chem. Int. Ed. 2001, 40, 2004–2021. [Google Scholar] [CrossRef]
- McKay, C.S.; Finn, M.G. Click chemistry in complex mixtures: Bioorthogonal bioconjugation. Chem. Biol. 2014, 21, 1075–1101. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Chao, C.S.; Li, C.W.; Chen, M.C.; Chang, S.S.; Mong, K.K. Low-concentration 1,2-trans beta-selective glycosylation strategy and its applications in oligosaccharide synthesis. Chem. Eur. J. 2009, 15, 10972–10982. [Google Scholar] [CrossRef] [Scilit]
- Mong, K.T.; Chao, C.; Chen, M.; Lin, C. Tandem one-pot acetalation-acetylation for direct access to differentially protected thioglycosides and O-glycosides with p-toluenesulfonic acid. Synlett 2009, 2009, 603–606. [Google Scholar] [CrossRef] [Scilit]
- D’Angelo, K.A.; Taylor, M.S. Borinic acid catalyzed stereo- and regioselective couplings of glycosyl methanesulfonates. J. Am. Chem. Soc. 2016, 138, 11058–11066. [Google Scholar] [CrossRef] [Scilit]
- Larsen, K.; Olsen, C.E.; Motawia, M.S. A facile protocol for direct conversion of unprotected sugars into phenyl 4,6-O-benzylidene-per-O-acetylated-1,2-trans-thioglycosides. Carbohydr. Res. 2003, 338, 199–202. [Google Scholar] [CrossRef] [Scilit]
- Li, Q.; Jaiswal, M.; Rohokale, R.S.; Guo, Z. A diversity-oriented strategy for chemoenzymatic synthesis of glycosphingolipids and related derivatives. Org. Lett. 2020, 22, 8245–8249. [Google Scholar] [CrossRef] [Scilit]
- Rohokale, R.S.; Li, Q.; Guo, Z. A diversity-oriented strategy for chemical synthesis of glycosphingolipids: Synthesis of glycosphingolipid LcGg4 and its analogues and derivatives. J. Org. Chem. 2021, 86, 1633–1648. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Meek, S.J.; O’Brien, R.V.; Llaveria, J.; Schrock, R.R.; Hoveyda, A.H. Catalytic Z-selective olefin cross-metathesis for natural product synthesis. Nature 2011, 471, 461–466. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Hasegawa, A.; Morita, M.; Kojima, Y.; Ishida, H.; Kiso, M. Synthesis of cerebroside, lactosyl ceramide, and ganglioside GM3 analogs containing beta-thioglycosidically linked ceramide. Carbohydr. Res. 1991, 214, 43–53. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Albrecht, B.; Pütz, U.; Schwarzmann, G. Synthesis of fluorescent and radioactive analogues of two lactosylceramides and glucosylceramide containing beta-thioglycosidic bonds that are resistant to enzymatic degradation. Carbohydr. Res. 1995, 276, 289–308. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Miura, Y.; Arai, T.; Yamagata, T. Synthesis of amphiphilic lactosides that possess a lactosylceramide-mimicking N-acyl structure: Alternative universal substrates for endo-type glycosylceramidases. Carbohydr. Res. 1996, 289, 193–199. [Google Scholar] [CrossRef] [Scilit]
- Ohlsson, J.; Magnusson, G. Analogues of glycosphingolipids and glycerolipids suitable for conjugation to gold- and amino-functionalised surfaces. Tetrahedron 2000, 56, 9975–9984. [Google Scholar] [CrossRef] [Scilit]
- Zhang, Y.; Toyokuni, T.; Ruan, F.; Hakomori, S.I. A one pot synthesis of mono- and di-lactosyl sphingosines. Glycoconj. J. 2001, 18, 557–563. [Google Scholar] [CrossRef] [Scilit]
- Liu, Y.; Bittman, R. Synthesis of fluorescent lactosylceramide stereoisomers. Chem. Phys. Lipids 2006, 142, 58–69. [Google Scholar] [CrossRef] [Scilit]
- Liu, Y.; Ding, N.; Xiao, H.; Li, Y. Efficient syntheses of a series of glycosphingolipids with 1,2-trans-glycosidic linkages. J. Carbohydr. Chem. 2006, 25, 471–489. [Google Scholar] [CrossRef] [Scilit]
- Morales-Serna, J.A.; Boutureira, O.; Díaz, Y.; Matheu, M.I.; Castillón, S. Highly efficient and stereoselective synthesis of beta-glycolipids. Org. Biomol. Chem. 2008, 6, 443–446. [Google Scholar] [CrossRef] [Scilit]
- Morales-Serna, J.A.; Díaz, Y.; Matheu, M.I.; Castillón, S. Efficient synthesis of β-glycosphingolipids by reaction of stannylceramides with glycosyl Iodides promoted by TBAI/AW 300 molecular sieves. Eur. J. Org. Chem. 2009, 2009, 3849–3852. [Google Scholar] [CrossRef] [Scilit]
- Tanaka, H.N.; Yoshimura, Y.; Dovichi, N.J.; Palcic, M.M.; Hindsgaul, O. A concise chemical synthesis of a fluorescent βGal-(1,4)-S-βGlc-Cer derivative and its enzymatic elongation by glycosyltransferases. Tetrahedron Lett. 2012, 53, 1812–1815. [Google Scholar] [CrossRef] [Scilit]
- Nagatsuka, T.; Uzawa, H.; Sato, K.; Kondo, S.; Izumi, M.; Yokoyama, K.; Ohsawa, I.; Seto, Y.; Neri, P.; Mori, H.; et al. Localized surface plasmon resonance detection of biological toxins using cell surface oligosaccharides on glyco chips. ACS Appl. Mater. Interfaces 2013, 5, 4173–4180. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Cheng, J.M.; Dangerfield, E.M.; Timmer, M.S.; Stocker, B.L. A divergent approach to the synthesis of iGb3 sugar and lipid analogues via a lactosyl 2-azido-sphingosine intermediate. Org. Biomol. Chem. 2014, 12, 2729–2736. [Google Scholar] [CrossRef] [Scilit]
- Son, S.H.; Daikoku, S.; Ohtake, A.; Suzuki, K.; Kabayama, K.; Ito, Y.; Kanie, O. Syntheses of lactosyl ceramide analogues carrying novel bifunctional BODIPY dyes directed towards the differential analysis of multiplexed glycosphingolipids by MS/MS using iTRAQ. Chem. Commun. 2014, 50, 3010–3013. [Google Scholar] [CrossRef] [Scilit]
- D’Angelo, K.A.; Taylor, M.S. Borinic acid-catalyzed stereo- and site-selective synthesis of β-glycosylceramides. Chem. Commun. 2017, 53, 5978–5980. [Google Scholar] [CrossRef] [Scilit]
Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2025 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license (https://creativecommons.org/licenses/by/4.0/).