
Phylogeographic Structure and Molecular Evolution of Squash Leaf Curl China Virus

 ,

 , ,
Abstract
1. Introduction
2. Materials and Methods
2.1. Collection and Alignment of SLCCNV Strain Sequences
2.2. Recombination Signal Detection
2.3. Phylogenetic Analysis of SLCCNV Strains
2.4. Selection Pressure and Neutrality Test Analysis
2.5. Population Genetic Parameter Calculation
3. Results
3.1. Recombination Patterns Were Observed in the SLCCNV Strains
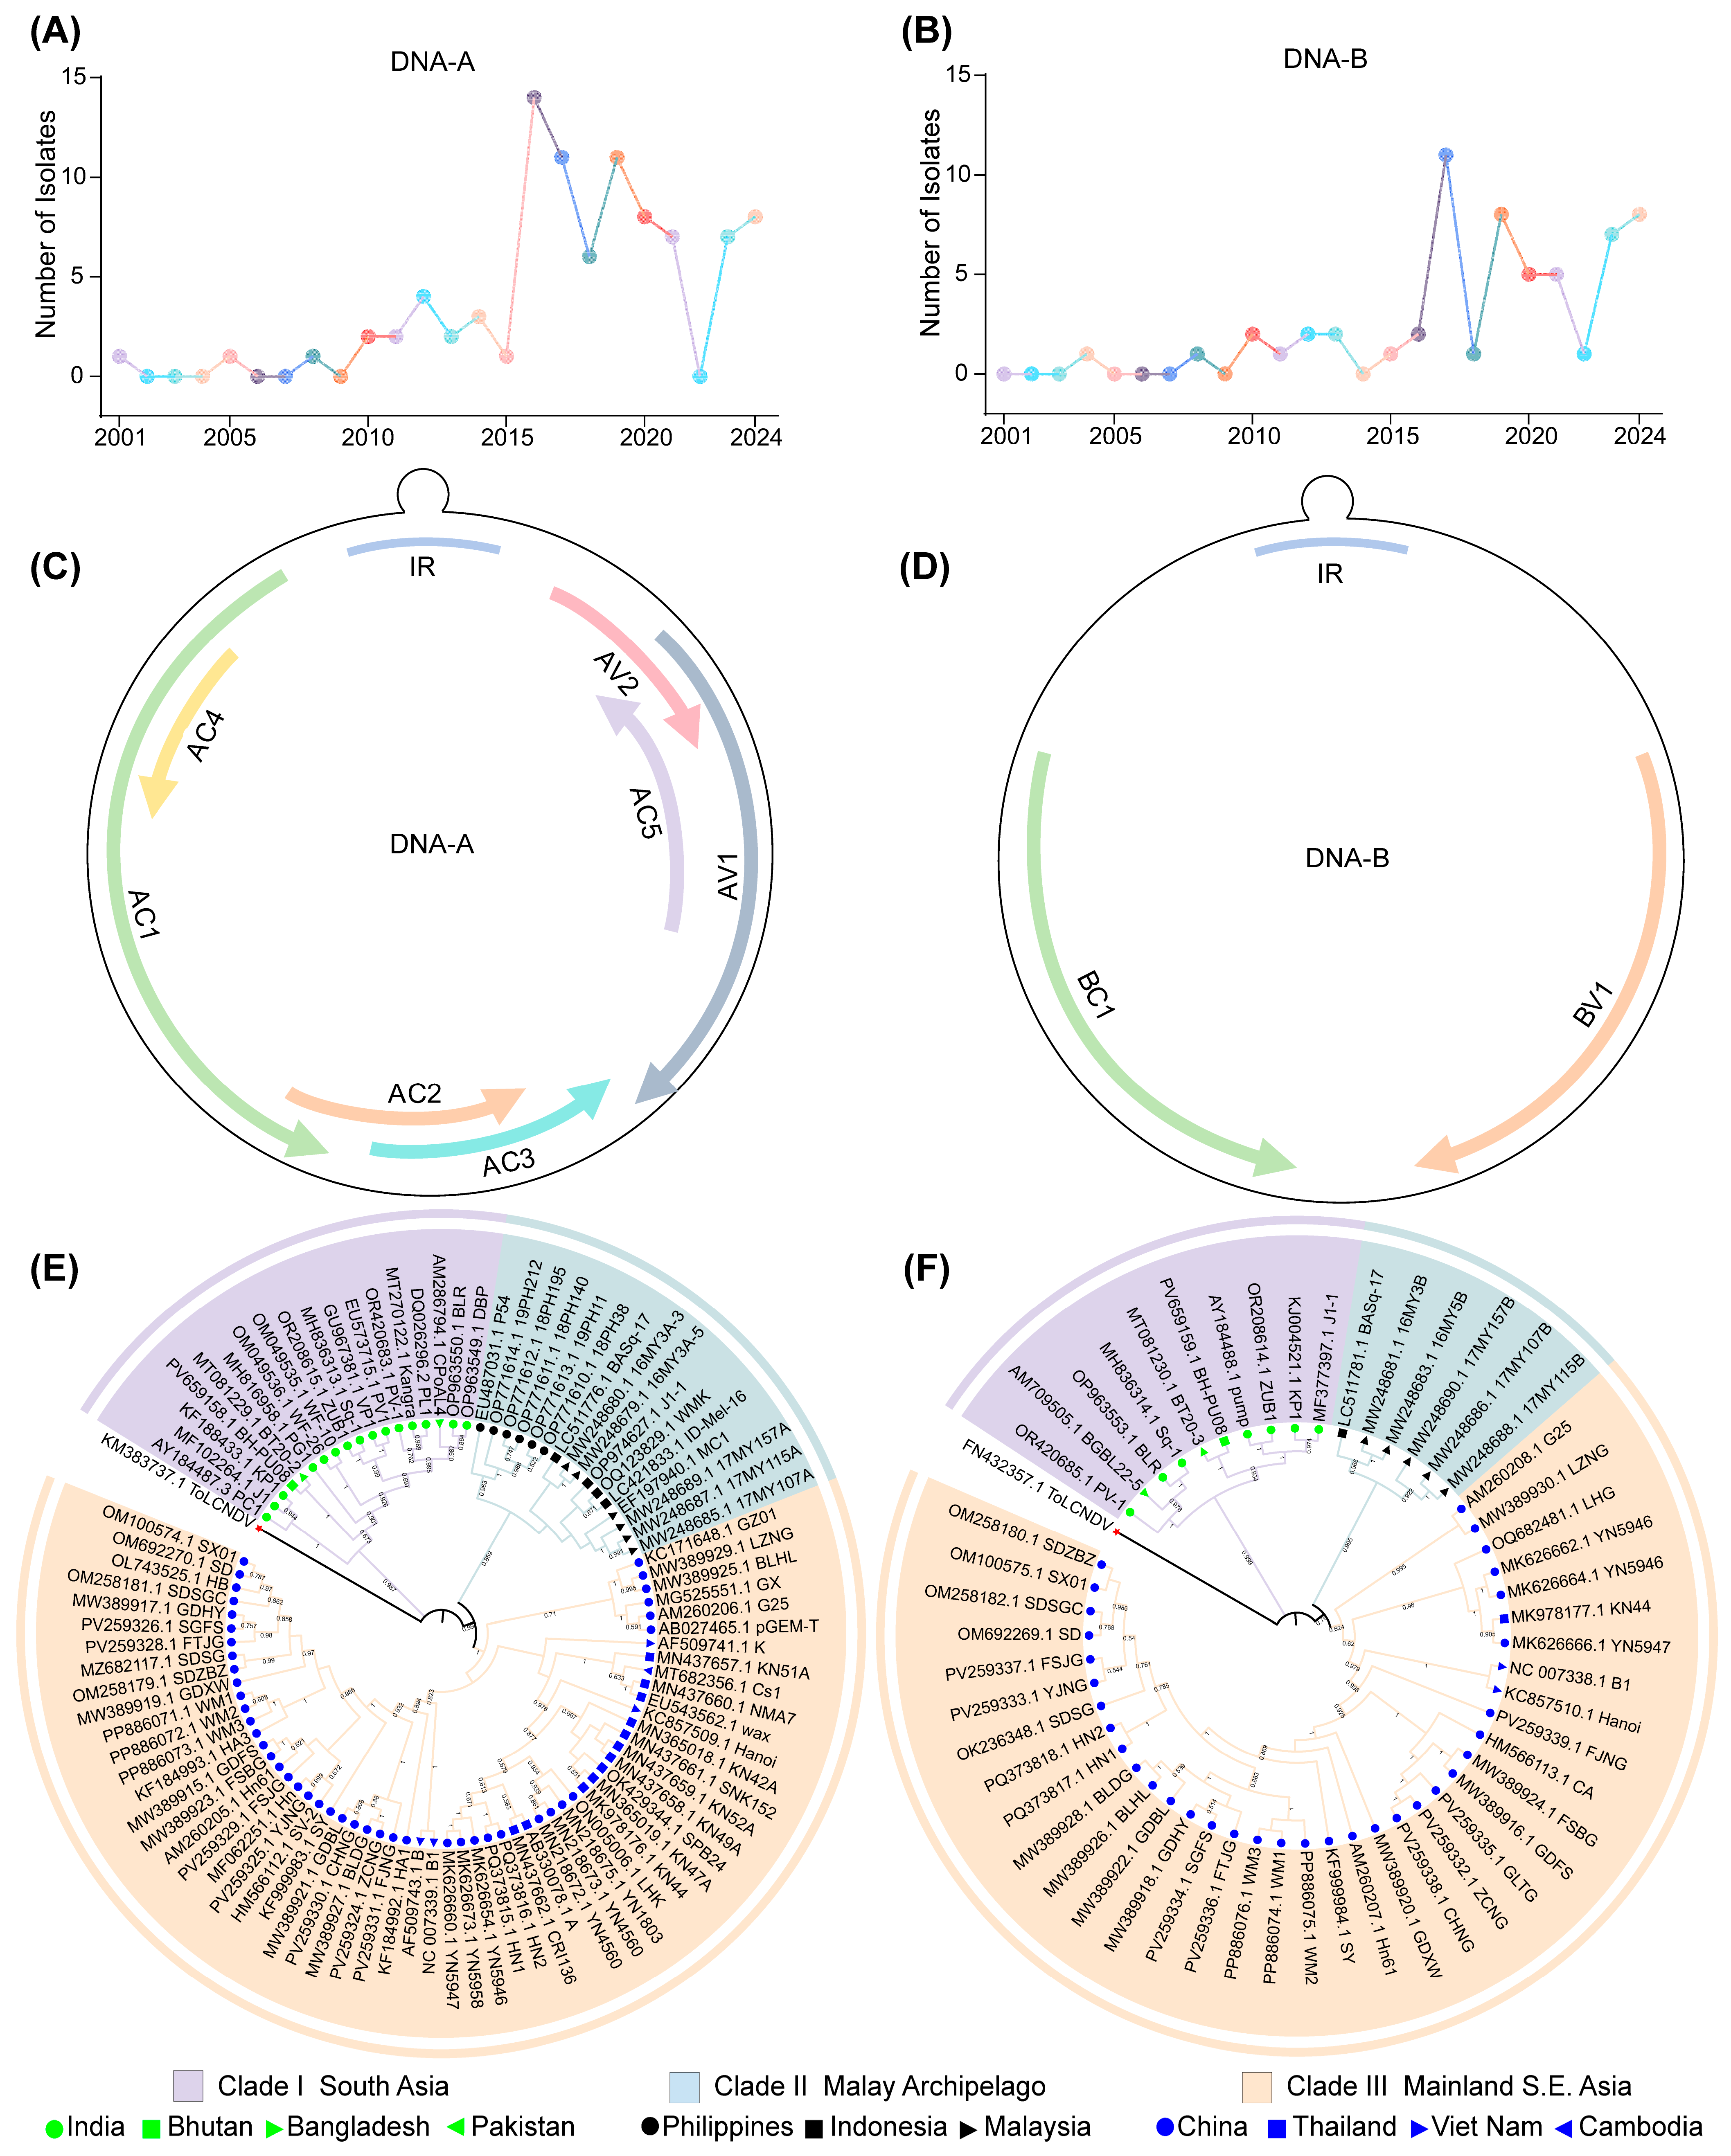
3.2. SLCCNV Strains Cluster into Three Genetic Groups
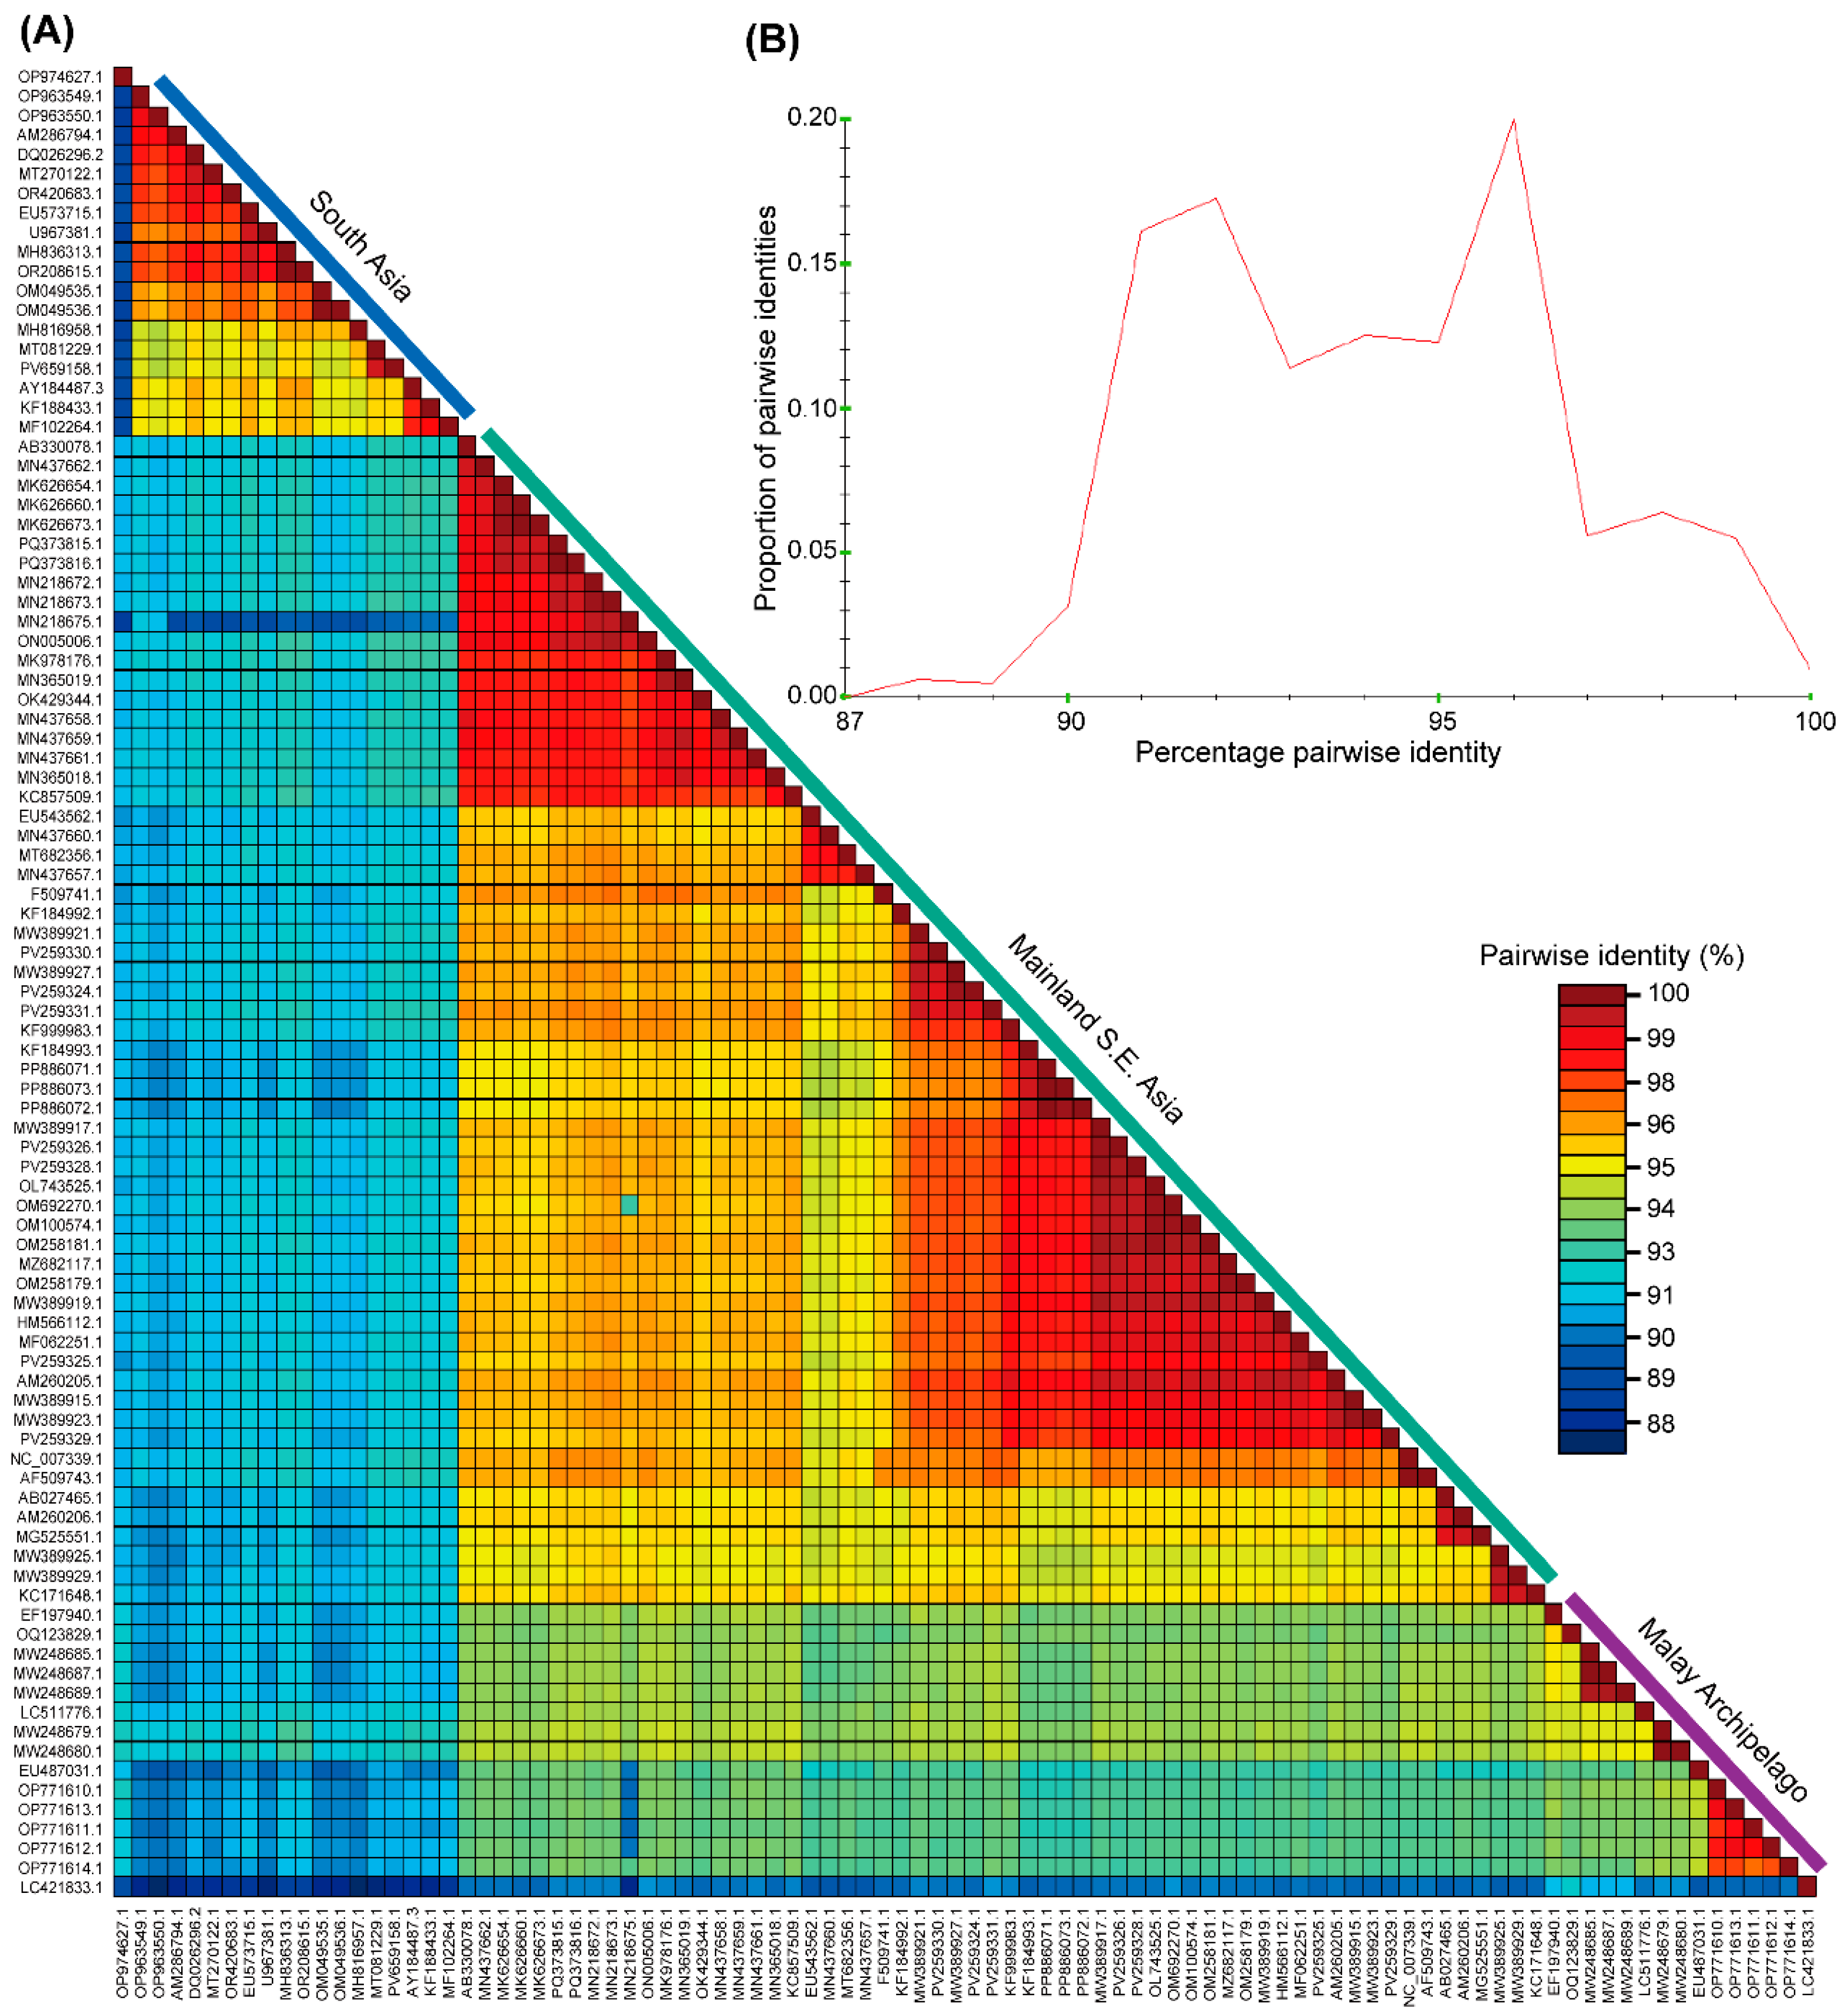
3.3. Sequence Identity Analysis in SLCCNV Strains
3.4. Analysis of Selection Pressure and Neutrality on SLCCNV Encoded Proteins
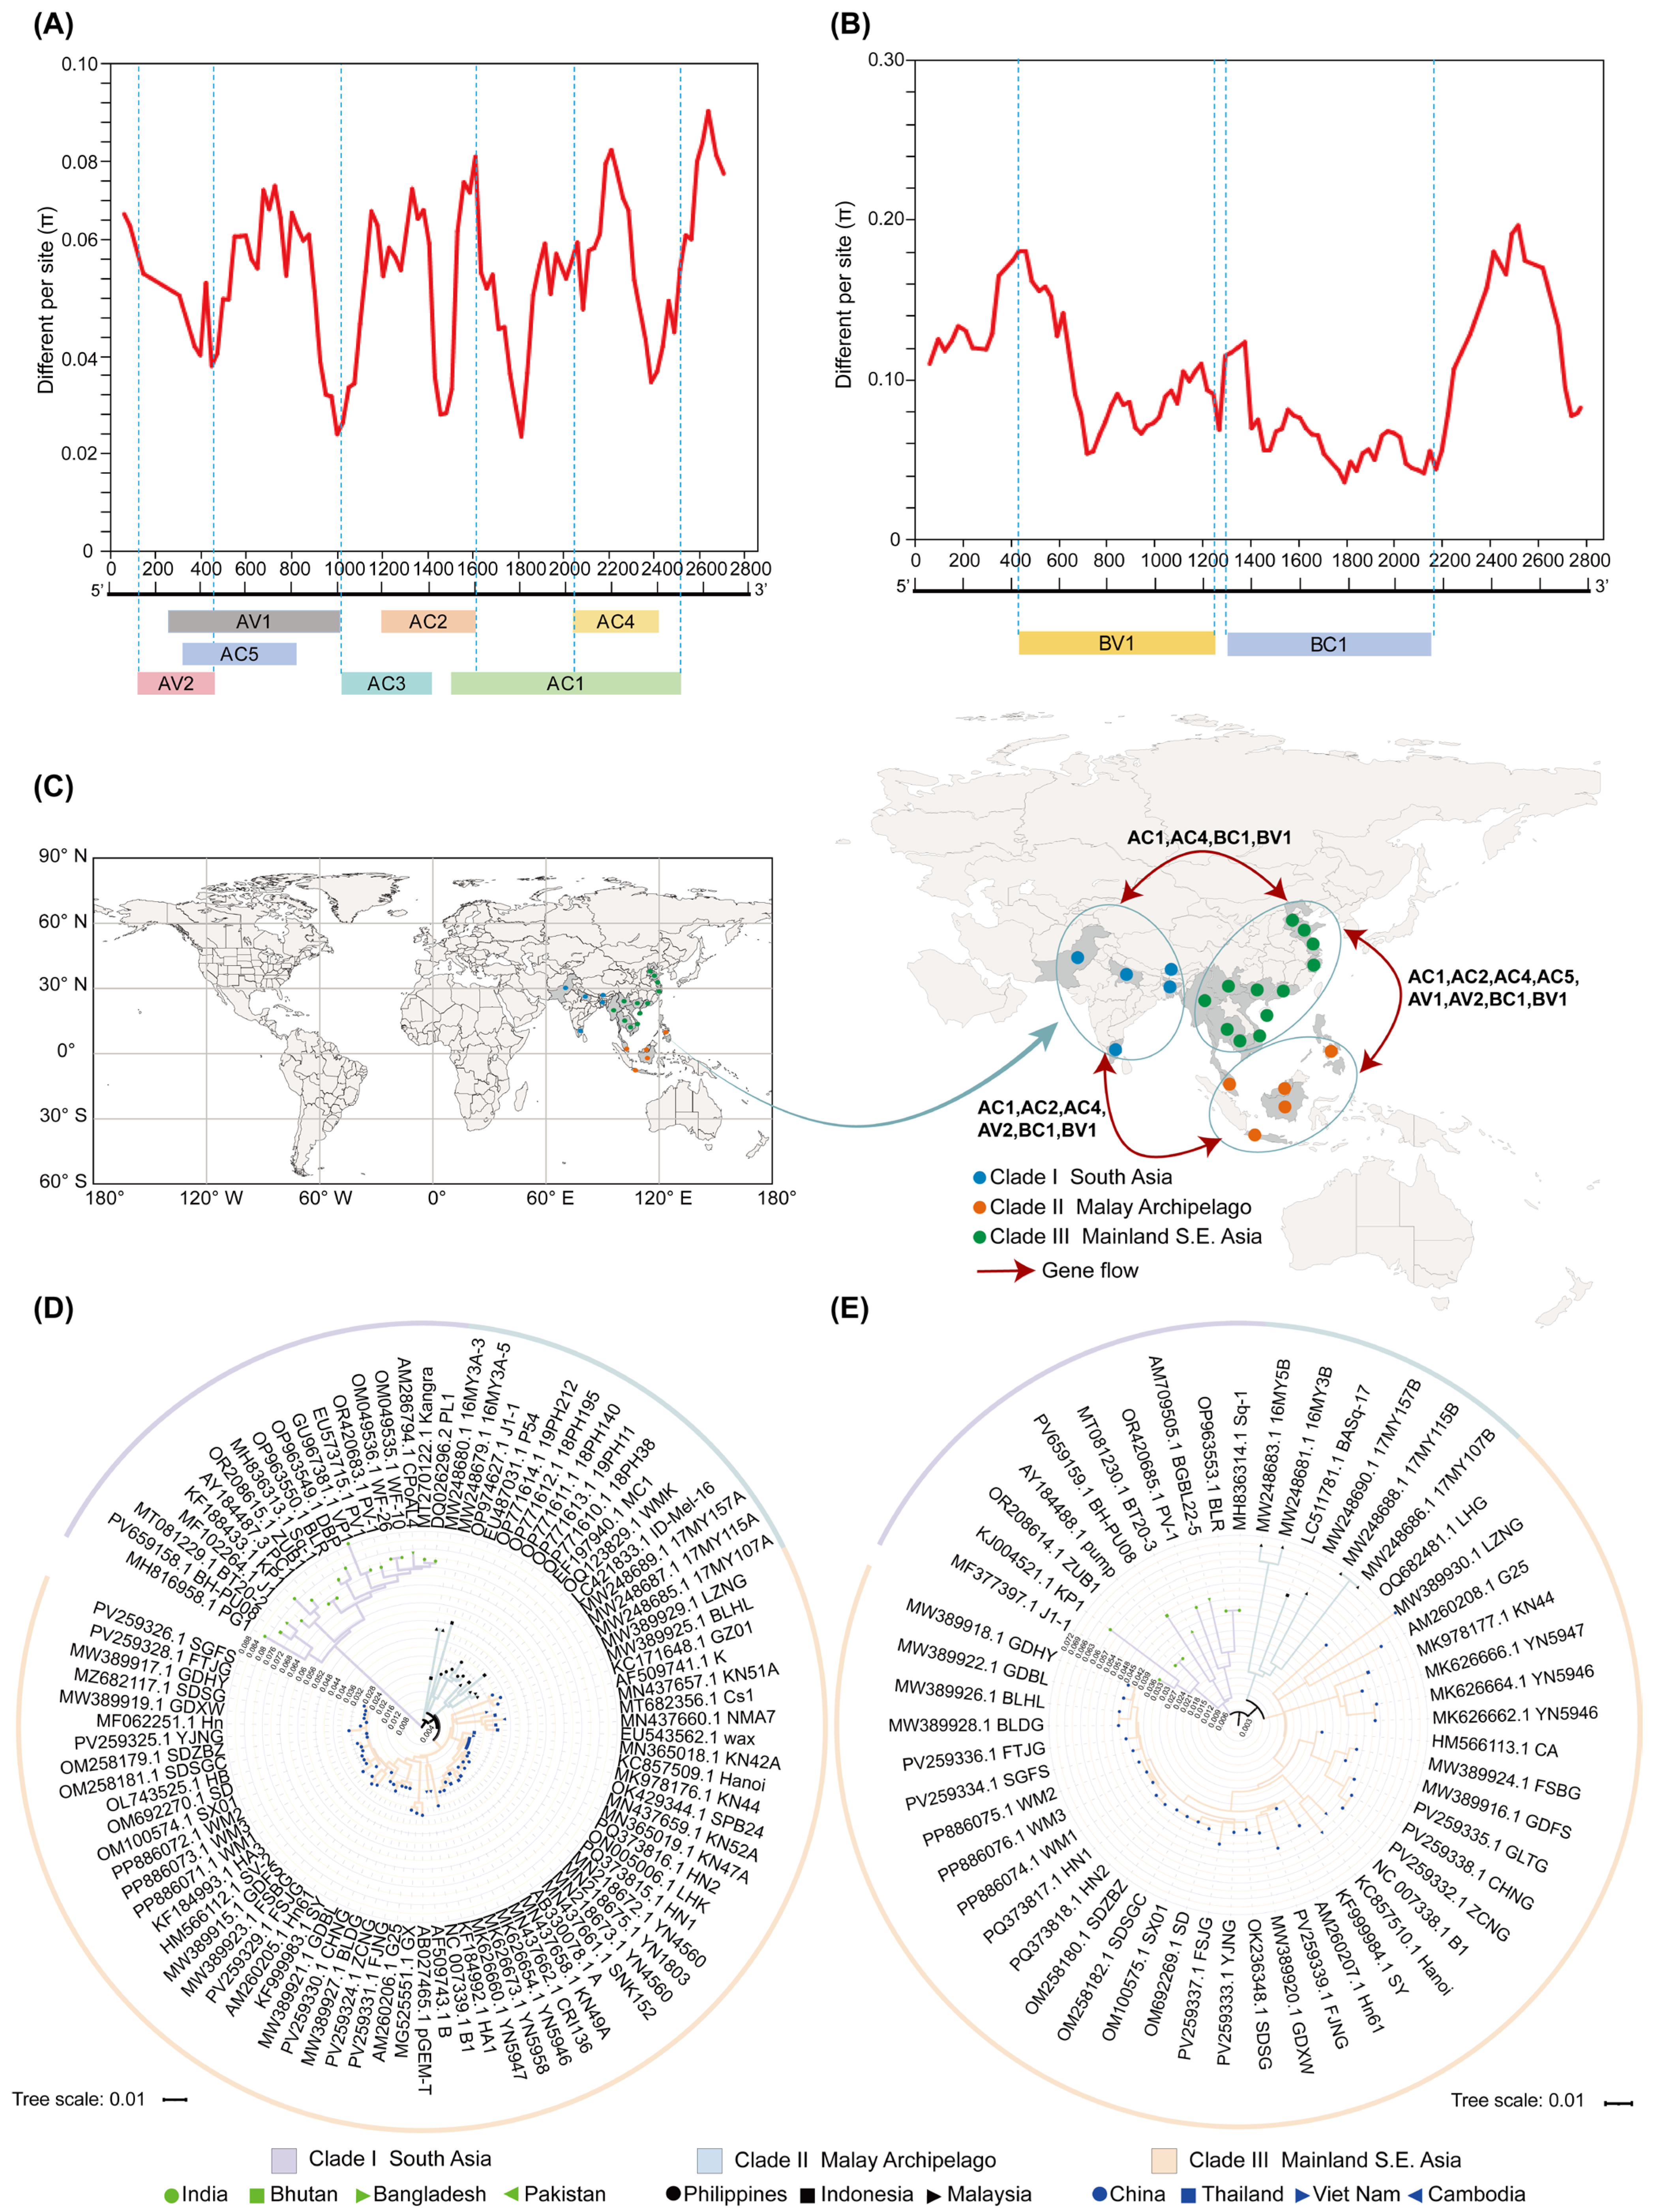
3.5. Heterogeneous Genetic Differentiation of SLCCNV Across Geographic Regions
4. Discussion
5. Conclusions
Supplementary Materials
Author Contributions
Funding
Data Availability Statement
Conflicts of Interest
References
- Polston, J.E.; De Barro, P.; Boykin, L.M. Transmission specificities of plant viruses with the newly identified species of the Bemisia tabaci species complex. Pest Manag. Sci. 2014, 70, 1547–1552. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Li, F.; Qiao, R.; Wang, Z.; Yang, X.; Zhou, X. Occurrence and distribution of geminiviruses in China. Sci. China Life Sci. 2022, 65, 1498–1503. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Venkataravanappa, V.; Kumar, H.D.V.; Nandan, M.; Hiremath, S.; Shankarappa, K.S.; Reddy, M.K.; Reddy, C.N.L. Characterization of squash leaf curl China virus associated with mosaic and vein clearing disease of pointed gourd (Trichosanthes dioica Roxb.) and its vector whitefly cryptic species. Indian Phytopathol. 2022, 75, 573–581. [Google Scholar] [CrossRef] [Scilit]
- Wu, H.; Li, M.; Hong, N.; Peng, B.; Gu, Q. Molecular and biological characterization of melon-infecting squash leaf curl China virus in China. J. Integr. Agric. 2020, 19, 570–577. [Google Scholar] [CrossRef] [Scilit]
- Ismayil, A.; Haxim, Y.; Wang, Y.; Li, H.; Qian, L.; Han, T.; Chen, T.; Jia, Q.; Liu, A.Y.; Zhu, S.; et al. Cotton leaf curl multan virus C4 protein suppresses both transcriptional and post-transcriptional gene silencing by interacting with SAM synthetase. PLoS Pathog. 2018, 14, e1007282. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Fondong, V.N. Geminivirus protein structure and function. Mol. Plant Pathol. 2013, 14, 635–649. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Wu, H.; Liu, M.; Kang, B.; Liu, L.; Hong, N.; Peng, B.; Gu, Q. AC5 protein encoded by squash leaf curl China virus is an RNA silencing suppressor and a virulence determinant. Front. Microbiol. 2022, 13, 980147. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Liu, L.; Wang, Y.; Geng, Y.; Yu, B.; Yan, L.; Hao, F.; Wu, H.; Wang, P.; Gu, Q.; Kang, B. Molecular characterization and pathogenicity of watermelon isolates of Begomovirus cucurbitachinaense. Int. J. Mol. Sci. 2025, 26, 4289. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Baldodiya, G.M.; Devi, K.; Borah, B.K.; Nath, P.D.; Modi, M.K. Characterization and in silico proteomic analysis of C2 and C3 proteins of squash leaf curl China virus associated with pumpkin leaf curl disease in Assam, India. Acta Virol. 2019, 63, 139–148. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Kumar, R.; Dasgupta, I. Geminiviral C4/AC4 proteins: An emerging component of the viral arsenal against plant defence. Virology 2023, 579, 156–168. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Padidam, M.; Beachy, R.N.; Fauquet, C.M. The role of AV2 (“precoat”) and coat protein in viral replication and movement in tomato leaf curl geminivirus. Virology 1996, 224, 390–404. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Schaffer, R.L.; Miller, C.G.; Petty, I.T. Virus and host-specific adaptations in the BL1 and BR1 genes of bipartite geminiviruses. Virology 1995, 214, 330–338. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Tahir, M.; Haider, M.S.; Briddon, R.W. First report of squash leaf curl China virus in Pakistan. Australas. Plant Dis. Notes 2010, 5, 21–24. [Google Scholar] [CrossRef] [Scilit]
- Saritha, R.K.; Bag, T.; Loganathan, M.; Rai, A.; Rai, M. First report of squash leaf curl China virus causing mosaic symptoms on summer squash (Cucurbita pepo) grown in Varanasi district of India. Arch. Phytopathol. Plant Prot. 2011, 44, 179–185. [Google Scholar] [CrossRef] [Scilit]
- Ito, T.; Ogawa, T.; Samretwanich, K.; Sharma, P.; Ikegami, M. Yellow leaf curl disease of pumpkin in Thailand is associated with squash leaf curl China virus. Plant Pathol. 2008, 57, 766. [Google Scholar] [CrossRef] [Scilit]
- Singh, R.; Raj, S.K.; Prasad, V. Molecular characterization of a strain of squash leaf curl China virus from north India. J. Phytopathol. 2008, 156, 222–228. [Google Scholar] [CrossRef] [Scilit]
- Riyaz, S.U.M.; Deepan, S.; Jesse, M.i.; Dharanivasan, G.; Kathiravan, K. New record of bipartite squash leaf curl China virus (SLCCNV) and croton yellow vein mosaic beta satellite associated with yellow vein disease of ash gourd in India. New Dis. Rep. 2015, 31, 3. [Google Scholar] [CrossRef] [Scilit]
- Riyaz, S.U.M.; Deepan, S.; Dharanivasan, G.; Jesse, M.I.; Muthuramalingam, R.; Kathiravan, K. First report on a variant of squash leaf curl China virus (SLCCNV) infecting Benincasa hispida in India. New Dis. Rep. 2013, 28, 20. [Google Scholar] [CrossRef] [Scilit]
- Yin, Q.; Yang, H.; Gong, Q.; Wang, H.; Liu, Y.; Hong, Y.; Tien, P. Tomato yellow leaf curl China virus: Monopartite genome organization and agroinfection of plants. Virus Res. 2001, 81, 69–76. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Zhou, X.P.; Xie, Y.; Zhang, Z.K. Molecular characterization of a distinct begomovirus infecting tobacco in yunnan, China. Arch. Virol. 2001, 146, 1599–1606. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Xie, Y.; Zhou, X.P. Molecular characterization of squash leaf curl yunnan virus, a new begomovirus and evidence for recombination. Arch. Virol. 2003, 148, 2047–2054. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- García-Arenal, F.; Fraile, A.; Malpica, J.M. Variability and genetic structure of plant virus populations. Annu. Rev. Phytopathol. 2001, 39, 157–186. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Yu, J.; Xiong, Y.; Li, Y.; Zhao, M.; Yang, X.; Wang, Y.; Huang, H.; Zhou, C.; Huang, W.; Qing, L. Genetic variation and molecular evolution of tomato yellow leaf curl China virus and its betasatellite DNA isolates in China. Phytopathol. Res. 2025, 7, 1–14. [Google Scholar] [CrossRef] [Scilit]
- Sun, S.-R.; Chen, J.-S.; He, E.-Q.; Huang, M.-T.; Fu, H.-Y.; Lu, J.-J.; Gao, S.-J. Genetic variability and molecular evolution of maize yellow mosaic virus populations from different geographic origins. Plant Dis. 2021, 105, 896–903. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Charoenvilaisiri, S.; Seepiban, C.; Phironrit, N.; Phuangrat, B.; Yoohat, K.; Deeto, R.; Chatchawankanphanich, O.; Gajanandana, O. Occurrence and distribution of begomoviruses infecting tomatoes, peppers and cucurbits in Thailand. Crop Prot. 2020, 127, 104948. [Google Scholar] [CrossRef] [Scilit]
- Du, J.; Li, S.; Yang, X.; Hao, S.; Li, J.; Tian, R.; Zhang, S.; Li, P. Molecular and biological characterization of squash leaf curl China virus infecting Phaseolus vulgaris in China. J. Plant Dis. Prot. 2025, 132, 104. [Google Scholar] [CrossRef] [Scilit]
- Kumar, S.; Stecher, G.; Suleski, M.; Sanderford, M.; Sharma, S.; Tamura, K. MEGA12: Molecular evolutionary genetic analysis version 12 for adaptive and green computing. Mol. Biol. Evol. 2024, 41, msae263. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Huson, D.H.; Bryant, D. Application of phylogenetic networks in evolutionary studies. Mol. Biol. Evol. 2006, 23, 254–267. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Martin, D.P.; Murrell, B.; Golden, M.; Khoosal, A.; Muhire, B. RDP4: Detection and analysis of recombination patterns in virus genomes. Virus Evol. 2015, 1, vev003. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Chinnaraja, C.; Viswanathan, R.; Karuppaiah, R.; Bagyalakshmi, K.; Malathi, P.; Parameswari, B. Complete genome characterization of sugarcane yellow leaf virus from India: Evidence for RNA recombination. Eur. J. Plant Pathol. 2013, 135, 335–349. [Google Scholar] [CrossRef] [Scilit]
- Lin, Y.-H.; Gao, S.-J.; Damaj, M.B.; Fu, H.-Y.; Chen, R.-K.; Mirkov, T.E. Genome characterization of sugarcane yellow leaf virus from China reveals a novel recombinant genotype. Arch. Virol. 2014, 159, 1421–1429. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Guo, M.; Li, M.; Liu, T.; Sun, W.; Du, K.; Yang, S.; Fu, Z.; Kou, Z. Epidemiological and genetic characteristics of sapovirus in Shandong, China, 2022–2023. Viruses 2025, 17, 469. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Tajima, F. The effect of change in population size on DNA polymorphism. Genetics 1989, 123, 597–601. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Rozas, J.; Ferrer-Mata, A.; Sánchez-DelBarrio, J.C.; Guirao-Rico, S.; Librado, P.; Ramos-Onsins, S.E.; Sánchez-Gracia, A. Dnasp 6: DNA sequence polymorphism analysis of large data sets. Mol. Biol. Evol. 2017, 34, 3299–3302. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Nei, M. Molecular Evolutionary Genetics; Columbia University Press: New York, NY, USA, 1987. [Google Scholar]
- Qiu, Y.; Zhang, H.; Tian, W.; Fan, L.; Du, M.; Yuan, G.; Wang, D.; Wen, C.; Xu, X. First report of squash leaf curl China virus infecting tomato in China. Plant Dis. 2022, 106, 2539. [Google Scholar] [CrossRef] [Scilit]
- Long, X.; Zhang, S.; Shen, J.; Du, Z.; Gao, F. Phylogeography and evolutionary dynamics of tobacco curly shoot virus. Viruses 2024, 16, 1850. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Li, M.; Li, C.; Jiang, K.; Li, K.; Zhang, J.; Sun, M.; Wu, G.; Qing, L. Characterization of pathogenicity-associated V2 protein of tobacco curly shoot virus. Int. J. Mol. Sci. 2021, 22, 923. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Rocha, E.S.D.O.; De Oliveira, J.G.; Dos Santos, J.R.; Rodrigues, G.O.L.; Figueiredo, L.B.; Pessanha, J.E.M.; Proietti, F.A.; Da Fonseca, F.G.; Bonjardim, C.A.; Ferreira, P.C.P.; et al. Recombinant envelope protein-based enzyme immunoassay for IgG antibodies is comparable to neutralization tests for epidemiological studies of dengue infection. J. Virol. Methods 2013, 187, 114–120. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Mar, T.B.; Xavier, C.A.D.; Lima, A.T.M.; Nogueira, A.M.; Silva, J.C.F.; Ramos-Sobrinho, R.; Lau, D.; Zerbini, F.M. Genetic variability and population structure of the New World begomovirus Euphorbia yellow mosaic virus. J. Gen. Virol. 2017, 98, 1537–1551. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Akram, M.; Kamaal, N.; Pratap, A.; Kumar, D.; Muin, A.; Sabale, P.R.; Aidbhavi, R.; Sunani, S.K.; Rathore, M.; Gupta, S.; et al. Exploring distribution and genomic diversity of begomoviruses associated with yellow mosaic disease of legume crops from India, highlighting the dominance of mungbean yellow mosaic India virus. Front. Microbiol. 2024, 15, 1451986. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Fiallo-Olivé, E.; Navas-Castillo, J. Begomoviruses: What is the secret(s) of their success? Trends Plant Sci. 2023, 28, 715–727. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Möller, S.R.; Maruthi, M.N. Viral and host factors involved in host gain and host loss by tomato leaf curl begomoviruses in tomato and cucumbers. Mol. Plant Pathol. 2026, 27, e70202. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Zhao, S.; Gong, P.; Liu, J.; Liu, H.; Lozano-Durán, R.; Zhou, X.; Li, F. Geminivirus C5 proteins mediate formation of virus complexes at plasmodesmata for viral intercellular movement. Plant Physiol. 2023, 193, 322–338. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Shivaprasad, P.V.; Akbergenov, R.; Trinks, D.; Rajeswaran, R.; Veluthambi, K.; Hohn, T.; Pooggin, M.M. Promoters, transcripts, and regulatory proteins of mungbean yellow mosaic geminivirus. J. Virol. 2005, 79, 8149–8163. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Rojas, M.R.; Noueiry, A.O.; Lucas, W.J.; Gilbertson, R.L. Bean dwarf mosaic geminivirus movement proteins recognize DNA in a form- and size-specific manner. Cell 1998, 95, 105–113. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Zhang, S.C.; Wege, C.; Jeske, H. Movement proteins (BC1 and BV1) of abutilon mosaic geminivirus are cotransported in and between cells of sink but not of source leaves as detected by green fluorescent protein tagging. Virology 2001, 290, 249–260. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Lefeuvre, P.; Martin, D.P.; Elena, S.F.; Shepherd, D.N.; Roumagnac, P.; Varsani, A. Evolution and ecology of plant viruses. Nat. Rev. Microbiol. 2019, 17, 632–644. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Navas-Castillo, J.; Fiallo-Olivé, E.; Sánchez-Campos, S. Emerging virus diseases transmitted by whiteflies. Annu. Rev. Phytopathol. 2011, 49, 219–248. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Rocha Carolina, S.; Castillo-Urquiza Gloria, P.; Lima Alison, T.M.; Silva Fábio, N.; Xavier Cesar, A.D.; Hora-Júnior Braz, T.; Beserra-Júnior José, E.A.; Malta Antonio, W.O.; Martin Darren, P.; Varsani, A.; et al. Brazilian begomovirus populations are highly recombinant, rapidly evolving, and segregated based on geographical location. J. Virol. 2013, 87, 5784–5799. [Google Scholar] [CrossRef] [Scilit] [PubMed]



| Putative Parent | Detection Method b | |||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Event | Putative Recombinant | Major | Minor | Breakpoint a | R | G | B | M | C | S | T | |
| DNA-A | 1 | J1 MN594504.1 | WMK OQ123829.1 | Unknown (SV-1) LC417095.1 | 56–1543 | ++ | ++ | ++ | ++ | ++ | ++ | ++ |
| 2 | SV-1 LC417095.1 | WMK OQ123829.1 | Unknown (PV1) EU573715.1 | 56–1544 | ++ | ++ | ++ | ++ | ++ | ++ | ++ | |
| 3 | Vir-6569 MK064240.1 | Cs1 MT682356.1 | Unknown (SX01) OM100574.1 | 998–1478 | ++ | ++ | ++ | ++ | ++ | ++ | ++ | |
| 4 | TMK OP963548.1 | PV1 EU573715.1 | Unknown (WMK) OQ123829.1 | 1473–1864 | ++ | ++ | ++ | ++ | + | + | ++ | |
| 5 | Pum JN587811.1 | Kangra MT270122.1 | Unknown (16MY5A) MW248682.1 | 1541–2543 | + | - | - | ++ | ++ | + | ++ | |
| 6 | 16MY5A MW248682.1 | PV-1 OR420683.1 | Unknown (PB1) OR135585.1 | 932–2564 | ++ | ++ | ++ | ++ | ++ | ++ | ++ | |
| 7 | WF-32 OM049537.1 | PV1 EU573715.1 | PG1 MH816957.1 | 1502–1980 | ++ | + | + | + | + | + | ++ | |
| DNA-B | 1 | PG1 MH816957.1 | DTMK OP963551.1 | Unknown (Varanasi) GU967382.1 | 1329–1925 | - | ++ | - | ++ | - | ++ | ++ |
| 2 | DTMK OP963551.1 | BLR OP963553.1 | GZ01 KC171649.1 | 710–1267 | ++ | ++ | ++ | ++ | ++ | ++ | ++ | |
| 3 | Pum JN624306.1 | BGBL22-5 AM709505.1 | ZUB1 OR208614.1 | 1333–2172 | ++ | ++ | ++ | ++ | ++ | ++ | ++ | |
| 4 | KM2 OR860426.1 | KP1 KJ004521.1 | Unknown (CPoBL2) AM778959.1 | 1–2498 | ++ | ++ | ++ | ++ | ++ | ++ | ++ | |
| 5 | Varanasi GU967382.1 | CPoBL2 AM778959.1 | BLR OP963553.1 | 63–2182 | ++ | ++ | ++ | ++ | - | ++ | ++ | |
| 6 | J1 MN594505.1 | J1-1 MF377397.1 | Unknown (BLR) OP963553.1 | 1968–2269 | ++ | ++ | - | ++ | ++ | + | ++ | |
| 7 | 17MY85B MW248684.1 | 17MY157B MW248690.1 | Unknown (GX2017) MG525552.1 | 28–2580 | ++ | ++ | ++ | + | ++ | - | ++ | |
| 8 | DBP OP963552.1 | CPoBL2 AM778959.1 | BLR OP963553.1 | 1535–2425 | - | - | ++ | ++ | - | ++ | ++ | |
| 9 | GX2017 MG525552.1 | LHG OQ682481.1 | FJNG PV259339.1 | 215–1505 | + | - | - | ++ | ++ | ++ | ++ | |
| 10 | GZ01 KC171649.1 | YN5947 MK626666.1 | GDFS MW389916.1 | 1210–1380 | - | - | - | ++ | ++ | + | + | |
| 11 | Cs1 MT682357.1 | Hanoi KC857510.1 | 17MY157B MW248690.1 | 346–1185 | - | - | - | + | ++ | ++ | + | |
| 12 | CPoBL2 AM778959.1 | BGBL22-5 AM709505.1 | DTMK OP963551.1 | 2150–2643 | ++ | + | ++ | + | + | ++ | + | |
| 13 | Varanasi-1 FJ859881.1 | BGBL22-5 AM709505.1 | SDSGC OM258182.1 | 2056–2122 | ++ | + | + | - | - | - | + | |
| 14 | Hn MF062252.1 | Hn61 AM260207.1 | B1 NC_007338.1 | 998–1602 | + | - | - | ++ | + | - | + | |
| Genomic Region a | Amino Acids (aa) | Nucleotide (nt) | Mutations b | ||||
|---|---|---|---|---|---|---|---|
| Length (aa) | ID (%) | InDels | Length (nt) | ID (%) | Syn | Non | |
| AC1 (Rep) | 238–373 | 90.74 | I, II | 717–1122 | 90.03 | 228 | 220 |
| AC2 (TrAP) | 133–135 | 91.94 | None | 402–408 | 95.50 | 43 | 121 |
| AC3 (Ren) | 104–137 | 94.39 | III | 351–414 | 95.24 | 70 | 91 |
| AC4 (SD) | 58 | 92.40 | None | 177 | 96.90 | 11 | 34 |
| AC5 (VSR) | 104–210 | 72.25 | IV | 315–633 | 77.11 | 21 | 170 |
| AV1 (CP) | 251–256 | 98.45 | V | 756–771 | 96.19 | 160 | 244 |
| AV2 (Pre-CP) | 111–161 | 74.86 | VI | 336–486 | 76.06 | 37 | 151 |
| BC1 (MP) | 269–305 | 87.86 | VII, VIII | 810–918 | 87.84 | 208 | 266 |
| BV1 (NSP) | 184–268 | 92.80 | None | 555–807 | 92.62 | 188 | 236 |
| Genomic Region a | Population | N b | Haplotype Diversity | Nucleotide Diversity | Tajima’s D c | dN/dS d |
|---|---|---|---|---|---|---|
| AC1 (Rep) | All (n = 94) | 80 | 0.996 ± 0.002 | 0.08997 ± 0.30602 | −2.41751 (**) | 0.25 |
| Clade I (n = 18) | 13 | 0.954 ± 0.034 | 0.12360 ± 0.25891 | −2.23190 (**) | 0.48 | |
| Clade II (n = 16) | 15 | 0.992 ± 0.025 | 0.06907 ± 0.09378 | −1.15208 (ns) | 0.18 | |
| Clade III (n = 60) | 53 | 0.995 ± 0.004 | 0.06371 ± 0.20009 | −2.43701 (**) | 0.25 | |
| AC2 (TrAP) | All (n = 94) | 72 | 0.993 ± 0.003 | 0.08286 ± 0.29108 | −2.43936 (**) | 0.74 |
| Clade I (n = 18) | 13 | 0.989 ± 0.031 | 0.02618 ± 0.03882 | −1.42770 (ns) | 0.67 | |
| Clade II (n = 16) | 17 | 0.982 ± 0.026 | 0.12220 ± 0.24839 | −2.14110 (**) | 0.94 | |
| Clade III (n = 60) | 44 | 0.988 ± 0.006 | 0.06457 ± 0.21444 | −2.48188 (**) | 0.76 | |
| AC3 (Ren) | All (n = 94) | 67 | 0.977 ± 0.009 | 0.06065 ± 0.12190 | −1.69596 (ns) | 0.32 |
| Clade I (n = 18) | 16 | 0.987 ± 0.023 | 0.03135 ± 0.05076 | −1.58043 (ns) | 0.53 | |
| Clade II (n = 16) | 13 | 0.981 ± 0.031 | 0.09051 ± 0.14891 | −1.73446 (ns) | 0.35 | |
| Clade III (n = 60) | 41 | 0.970 ± 0.013 | 0.02921 ± 0.04905 | −1.40344 (ns) | 0.27 | |
| AC4 (SD) | All (n = 94) | 55 | 0.981 ± 0.005 | 0.09963 ± 0.36444 | −2.46673 (**) | 1.12 |
| Clade I (n = 18) | 12 | 0.935 ± 0.041 | 0.12304 ± 0.25624 | −2.19926 (**) | 1.47 | |
| Clade II (n = 16) | 14 | 0.983 ± 0.028 | 0.04543 ± 0.06981 | −1.46725 (ns) | 1.17 | |
| Clade III (n = 60) | 29 | 0.959 ± 0.011 | 0.09950 ± 0.37991 | −2.62156 (ns) | 0.99 | |
| AC5 (VSR) | All (n = 94) | 64 | 0.985 ± 0.006 | 0.26409 ± 0.31843 | −0.58291 (ns) | 1.12 |
| Clade I (n = 18) | 16 | 0.987 ± 0.023 | 0.18486 ± 0.41808 | −2.38188 (**) | 1.06 | |
| Clade II (n = 16) | 14 | 0.983 ± 0.028 | 0.04078 ± 0.05362 | −1.03000 (ns) | 5.10 | |
| Clade III (n = 60) | 34 | 0.965 ± 0.013 | 0.03864 ± 0.05261 | −0.91627 (ns) | 3.61 | |
| AV1 (CP) | All (n = 94) | 74 | 0.994 ± 0.003 | 0.15241 ± 0.35818 | −1.96550 (*) | 0.41 |
| Clade I (n = 18) | 16 | 0.987 ± 0.023 | 0.09778 ± 0.23110 | −2.45806 (**) | 0.64 | |
| Clade II (n = 16) | 14 | 0.983 ± 0.028 | 0.03911 ± 0.05160 | −1.04920 (ns) | 0.06 | |
| Clade III (n = 60) | 45 | 0.988 ± 0.006 | 0.15836 ± 0.29448 | −1.65464 (ns) | 0.69 | |
| AV2 (Pre-CP) | All (n = 94) | 80 | 0.996 ± 0.002 | 0.52620 ± 0.53823 | −0.07658 (ns) | 0.91 |
| Clade I (n = 18) | 14 | 0.967 ± 0.030 | 0.18097 ± 0.40091 | −2.33945 (**) | 0.90 | |
| Clade II (n = 16) | 15 | 0.992 ± 0.025 | 0.28255 ± 0.52717 | −2.03394 (*) | 0.84 | |
| Clade III (n = 60) | 51 | 0.995 ± 0.004 | 0.53725 ± 0.51693 | 0.14091 (ns) | 0.96 | |
| BC1 (MP) | All (n = 53) | 44 | 0.993 ± 0.005 | 0.19036 ± 0.41487 | −1.96736 (*) | 0.50 |
| Clade I (n = 10) | 8 | 0.956 ± 0.059 | 0.05425 ± 0.06317 | −0.70244 (ns) | 0.07 | |
| Clade II (n = 6) | 5 | 0.933 ± 0.122 | 0.06769 ± 0.06833 | −0.06090 (ns) | 0.07 | |
| Clade III (n = 37) | 31 | 0.989 ± 0.009 | 0.22655 ± 0.43355 | −1.81424 (*) | 0.72 | |
| BV1 (NSP) | All (n = 53) | 46 | 0.994 ± 0.005 | 0.18359 ± 0.44231 | −2.12478 (*) | 0.64 |
| Clade I (n = 10) | 8 | 0.956 ± 0.059 | 0.18963 ± 0.29616 | −1.80196 (*) | 0.74 | |
| Clade II (n = 6) | 6 | 1.000 ± 0.096 | 0.31178 ± 0.39245 | −1.33843 (ns) | 0.54 | |
| Clade III (n = 37) | 32 | 0.991 ± 0.009 | 0.13025 ± 0.36601 | −2.44405 (**) | 0.25 |
| Genomic Region a | Comparisons | Ks * (p Value b) | Z * (p Value b) | Snn (p Value b) | Fst c | Nm d |
|---|---|---|---|---|---|---|
| AC1 (Rep) | Clade I (n = 18) versus Clade II (n = 16) | 3.50888 (0.0000 ***) | 4.85026 (0.0000 ***) | 0.97059 (0.0000 ***) | 0.19862 | 2.02 |
| Clade I (n = 18) versus Clade III (n = 60) | 3.29400 (0.0000 ***) | 6.62294 (0.0000 ***) | 0.97436 (0.0000 ***) | 0.22424 | 1.73 | |
| Clade II (n = 16) versus Clade III (n = 60) | 3.44938 (0.0000 ***) | 6.67794 (0.0000 ***) | 0.98684 (0.0000 ***) | 0.23808 | 1.60 | |
| AC2 (TrAP) | Clade I (n = 18) versus Clade II (n = 16) | 2.94546 (0.0000 ***) | 4.92823 (0.0000 ***) | 0.79293 (0.0030 **) | 0.19222 | 2.10 |
| Clade I (n = 18) versus Clade III (n = 60) | 2.39109 (0.0000 ***) | 6.50911 (0.0000 ***) | 1.00000 (0.0000 ***) | 0.45071 | 0.61 | |
| Clade II (n = 16) versus Clade III (n = 60) | 2.60453 (0.0000 ***) | 6.81203 (0.0000 ***) | 0.98734 (0.0000 ***) | 0.14292 | 3.00 | |
| AC3 (Ren) | Clade I (n = 18) versus Clade II (n = 16) | 2.56062 (0.0000 ***) | 4.73879 (0.0000 ***) | 1.00000 (0.0000 ***) | 0.36499 | 0.87 |
| Clade I (n = 18) versus Clade III (n = 60) | 2.13830 (0.0000 ***) | 6.60721 (0.0000 ***) | 1.00000 (0.0000 ***) | 0.63227 | 0.29 | |
| Clade II (n = 16) versus Clade III (n = 60) | 2.54404 (0.0000 ***) | 6.65855 (0.0000 ***) | 1.00000 (0.0000 ***) | 0.36704 | 0.86 | |
| AC4 (SD) | Clade I (n = 18) versus Clade II (n = 16) | 2.15290 (0.0000 ***) | 4.95924 (0.0000 ***) | 1.00000 (0.0000 ***) | 0.14240 | 3.01 |
| Clade I (n = 18) versus Clade III (n = 60) | 1.99564 (0.0000 ***) | 6.74982 (0.0000 ***) | 0.94338 (0.0000 ***) | 0.10171 | 4.42 | |
| Clade II (n = 16) versus Clade III (n = 60) | 2.15290 (0.0000 ***) | 4.95924 (0.0000 ***) | 1.00000 (0.0000 ***) | 0.14240 | 3.01 | |
| AC5 (VSR) | Clade I (n = 18) versus Clade II (n = 16) | 2.83643 (0.0000 ***) | 4.69172 (0.0000 ***) | 0.97059 (0.0000 ***) | 0.84108 | 0.09 |
| Clade I (n = 18) versus Clade III (n = 60) | 2.46647 (0.0000 ***) | 6.54332 (0.0000 ***) | 0.97835 (0.0000 ***) | 0.84526 | 0.09 | |
| Clade II (n = 16) versus Clade III (n = 60) | 2.40992 (0.0000 ***) | 6.68095 (0.0000 ***) | 1.00000 (0.0000 ***) | 0.30352 | 1.15 | |
| AV1 (CP) | Clade I (n = 18) versus Clade II (n = 16) | 2.63987 (0.0000 ***) | 4.68293 (0.0000 ***) | 0.97059 (0.0000 ***) | 0.50094 | 0.50 |
| Clade I (n = 18) versus Clade III (n = 60) | 2.90547 (0.0000 ***) | 6.61566 (0.0000 ***) | 0.98718 (0.0000 ***) | 0.36052 | 0.89 | |
| Clade II (n = 16) versus Clade III (n = 60) | 3.30675 (0.0000 ***) | 6.71370 (0.0000 ***) | 1.00000 (0.0000 ***) | 0.13991 | 3.07 | |
| AV2 (Pre-CP) | Clade I (n = 18) versus Clade II (n = 16) | 2.94193 (0.0000 ***) | 4.85203 (0.0000 ***) | 0.97059 (0.0000 ***) | 0.19735 | 2.03 |
| Clade I (n = 18) versus Clade III (n = 60) | 4.00309 (0.0000 ***) | 6.64058 (0.0000 ***) | 0.98701 (0.0000 ***) | 0.41618 | 0.70 | |
| Clade II (n = 16) versus Clade III (n = 60) | 4.35847 (0.0000 ***) | 6.74824 (0.0000 ***) | 0.93333 (0.0000 ***) | 0.30047 | 1.16 | |
| BC1 (MP) | Clade I (n = 10) versus Clade II (n = 6) | 3.72115 (0.0000 ***) | 3.22690 (0.0000 ***) | 1.00000 (0.0010 **) | 0.32739 | 1.03 |
| Clade I (n = 10) versus Clade III (n = 37) | 4.05877 (0.0000 ***) | 5.76297 (0.0000 ***) | 1.00000 (0.0000 ***) | 0.17802 | 2.31 | |
| Clade II (n = 6) versus Clade III (n = 37) | 4.11281 (0.0000 ***) | 5.65839 (0.0000 ***) | 0.98837 (0.0000 ***) | 0.17385 | 2.38 | |
| BV1 (NSP) | Clade I (n = 10) versus Clade II (n = 6) | 4.10627 (0.0000 ***) | 3.35998 (0.0000 ***) | 0.93750 (0.0020 **) | 0.09756 | 4.63 |
| Clade I (n = 10) versus Clade III (n = 37) | 3.56271 (0.0000 ***) | 5.69332 (0.0000 ***) | 0.95745 (0.0000 ***) | 0.25208 | 1.48 | |
| Clade II (n = 6) versus Clade III (n = 37) | 3.59772 (0.0000 ***) | 5.63883 (0.0000 ***) | 0.97674 (0.0000 ***) | 0.14109 | 3.04 |
Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2026 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license.
Share and Cite
Yu, J.; Han, X.; Liu, Y.; Peng, D.; Peng, H.; Kang, H.; Li, M.; Wu, G.; Qing, L.; Huang, W. Phylogeographic Structure and Molecular Evolution of Squash Leaf Curl China Virus. Viruses 2026, 18, 794. https://doi.org/10.3390/v18070794
Yu J, Han X, Liu Y, Peng D, Peng H, Kang H, Li M, Wu G, Qing L, Huang W. Phylogeographic Structure and Molecular Evolution of Squash Leaf Curl China Virus. Viruses. 2026; 18(7):794. https://doi.org/10.3390/v18070794
Chicago/Turabian StyleYu, Jingwen, Xue Han, Yaqin Liu, Deliang Peng, Huan Peng, Houxiang Kang, Mingjun Li, Gentu Wu, Ling Qing, and Wenkun Huang. 2026. "Phylogeographic Structure and Molecular Evolution of Squash Leaf Curl China Virus" Viruses 18, no. 7: 794. https://doi.org/10.3390/v18070794
APA StyleYu, J., Han, X., Liu, Y., Peng, D., Peng, H., Kang, H., Li, M., Wu, G., Qing, L., & Huang, W. (2026). Phylogeographic Structure and Molecular Evolution of Squash Leaf Curl China Virus. Viruses, 18(7), 794. https://doi.org/10.3390/v18070794