


Abstract
Based on the diaryl hydroxylamine scaffold, which exhibits the potential to inhibit all three enzymes of the first step of the kynurenine pathway, the main tryptophan degradation pathway in mammals, which is often activated in cancer, we report herein the synthesis of a ferrocenyl analogue as an attempt to improve the scaffold’s pan-inhibitory potency through the isosteric replacement of a phenyl group with the ferrocenyl moiety. The synthetic methodology followed gives access to O-((ferrocenyl)(aryl)methyl)hydroxylamines, a class of compounds not yet reported in the literature.
1. Introduction
The use of ferrocene building blocks in medicinal chemistry is an intriguing research area. Apart from its anticancer, antibacterial, antifungal, and antiparasitic potential [1], the ferrocenyl moiety has been increasingly used in drug design [2]. In particular, the bioisosteric replacement of a phenyl ring or heteroaromatic with the ferrocenyl group can lead to compounds with enhanced potency and/or selectivity, compared with the parent, purely organic molecule [3].
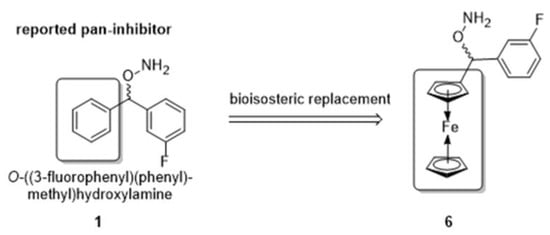
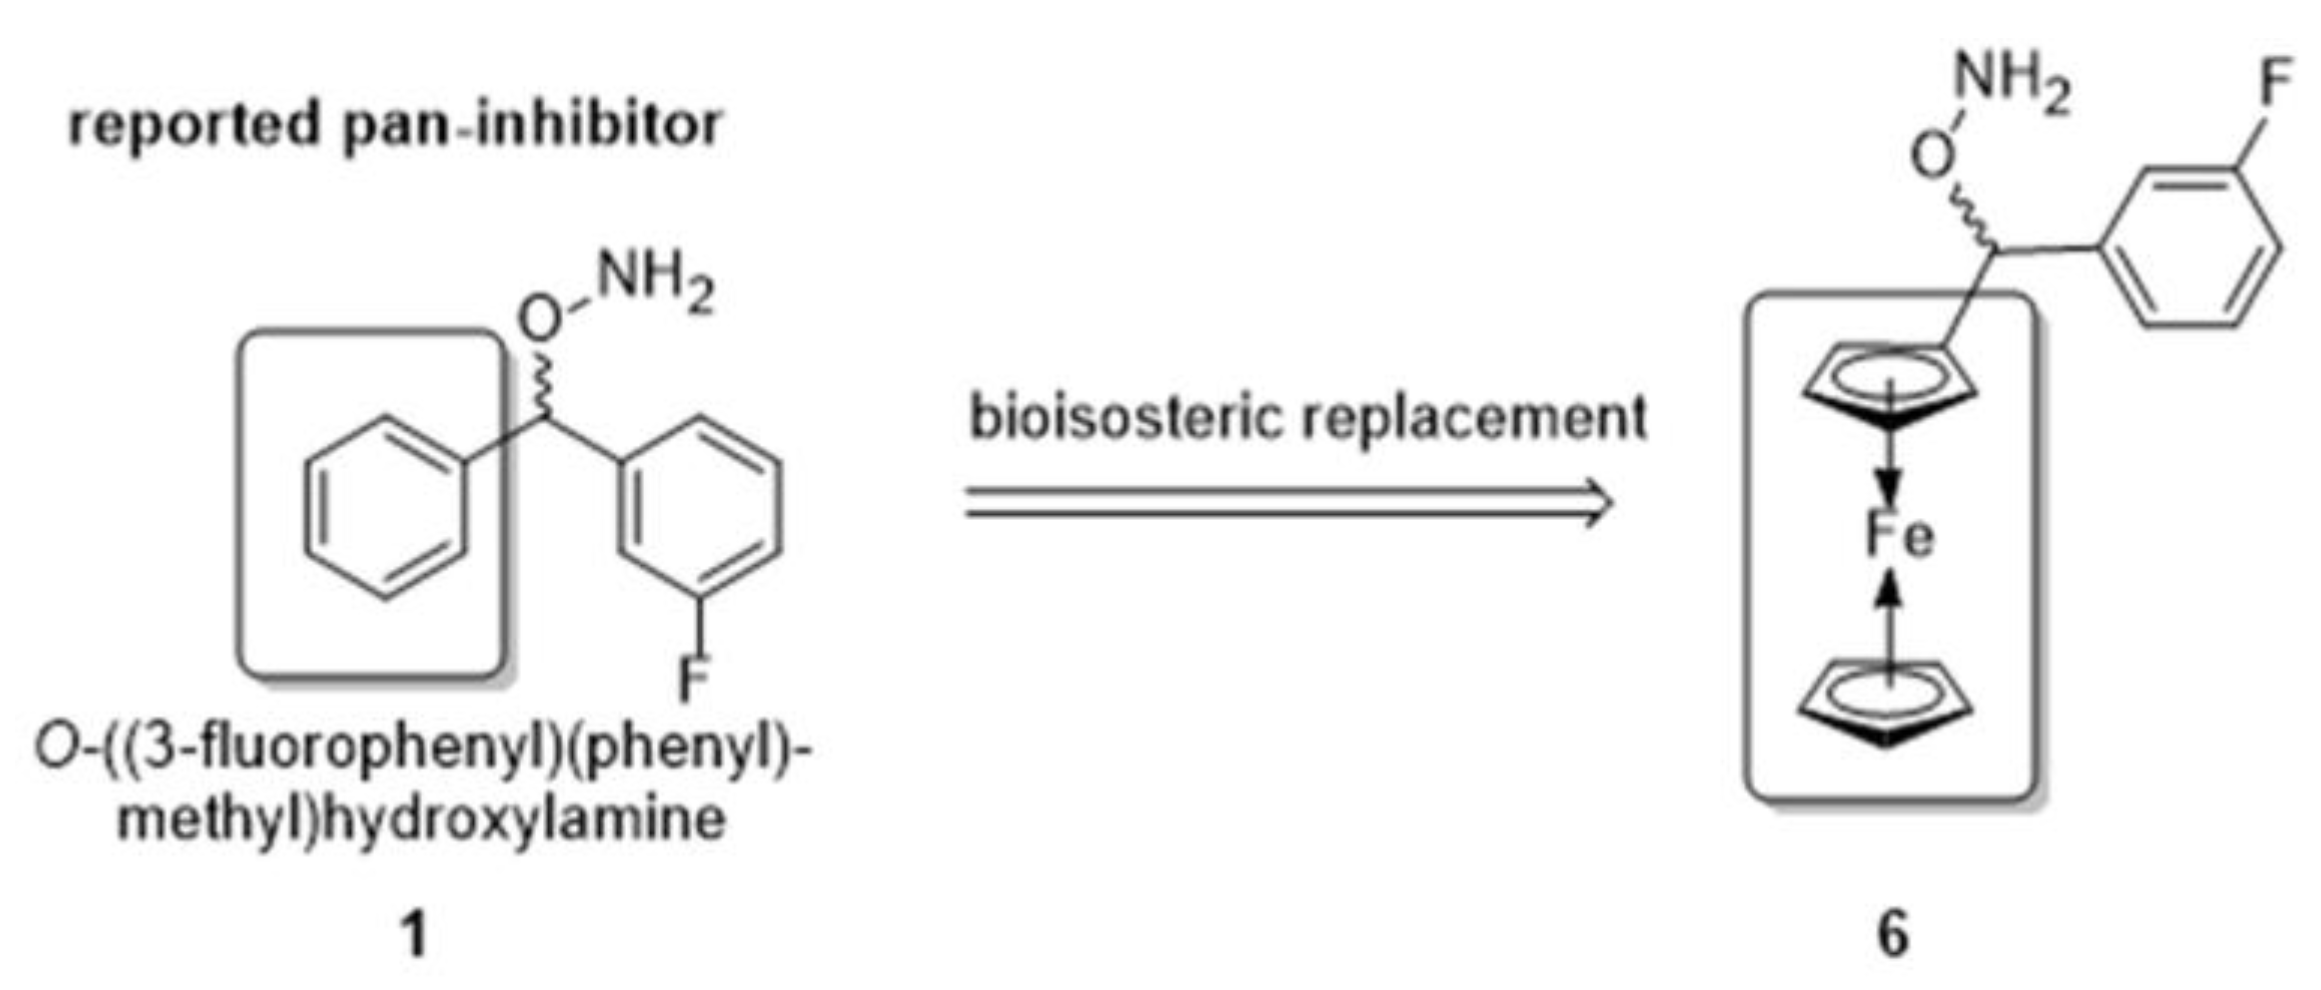
The diaryl hydroxylamine scaffold was recently identified as promising in its ability to pan-inhibit three enzymes that catalyze the first and rate-limiting step of the kynurenine pathway (KP), the major route of tryptophan degradation in vertebrates that is often activated in cancer. Diaryl hydroxylamines with a wide variety of substituted phenyl groups were evaluated [4,5] and O-((3-fluorophenyl)(phenyl)methyl)hydroxylamine 1 (Figure 1) was identified as one of the best analogues but still not with the desired potency.
Figure 1.
Bioisosteric replacement of a phenyl group with ferrocenyl moiety on a known KP pan-inhibitor.
As part of a medicinal chemistry program in our laboratory aiming to develop dual and/or pan-inhibitors of the KP, we present here the synthesis of the title compound O-((ferrocenyl)(3-fluorophenyl)methyl)hydroxylamine, in which the phenyl group of 1 was substituted with the ferrocenyl moiety (6, Figure 1) in an attempt to increase the scaffold’s potency for pan-inhibition. To the best of our knowledge, this is the first time a metallocene analogue of diaryl O-alkylhydroxylamines is reported. Despite the simplicity of its structure, the synthesis of 6 was accompanied by some difficulties, which were successfully overcome, thus paving the way toward the synthesis of such compounds.
2. Results and Discussion
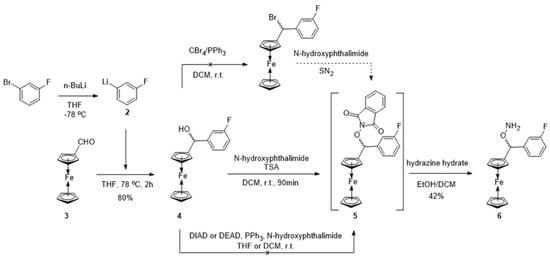
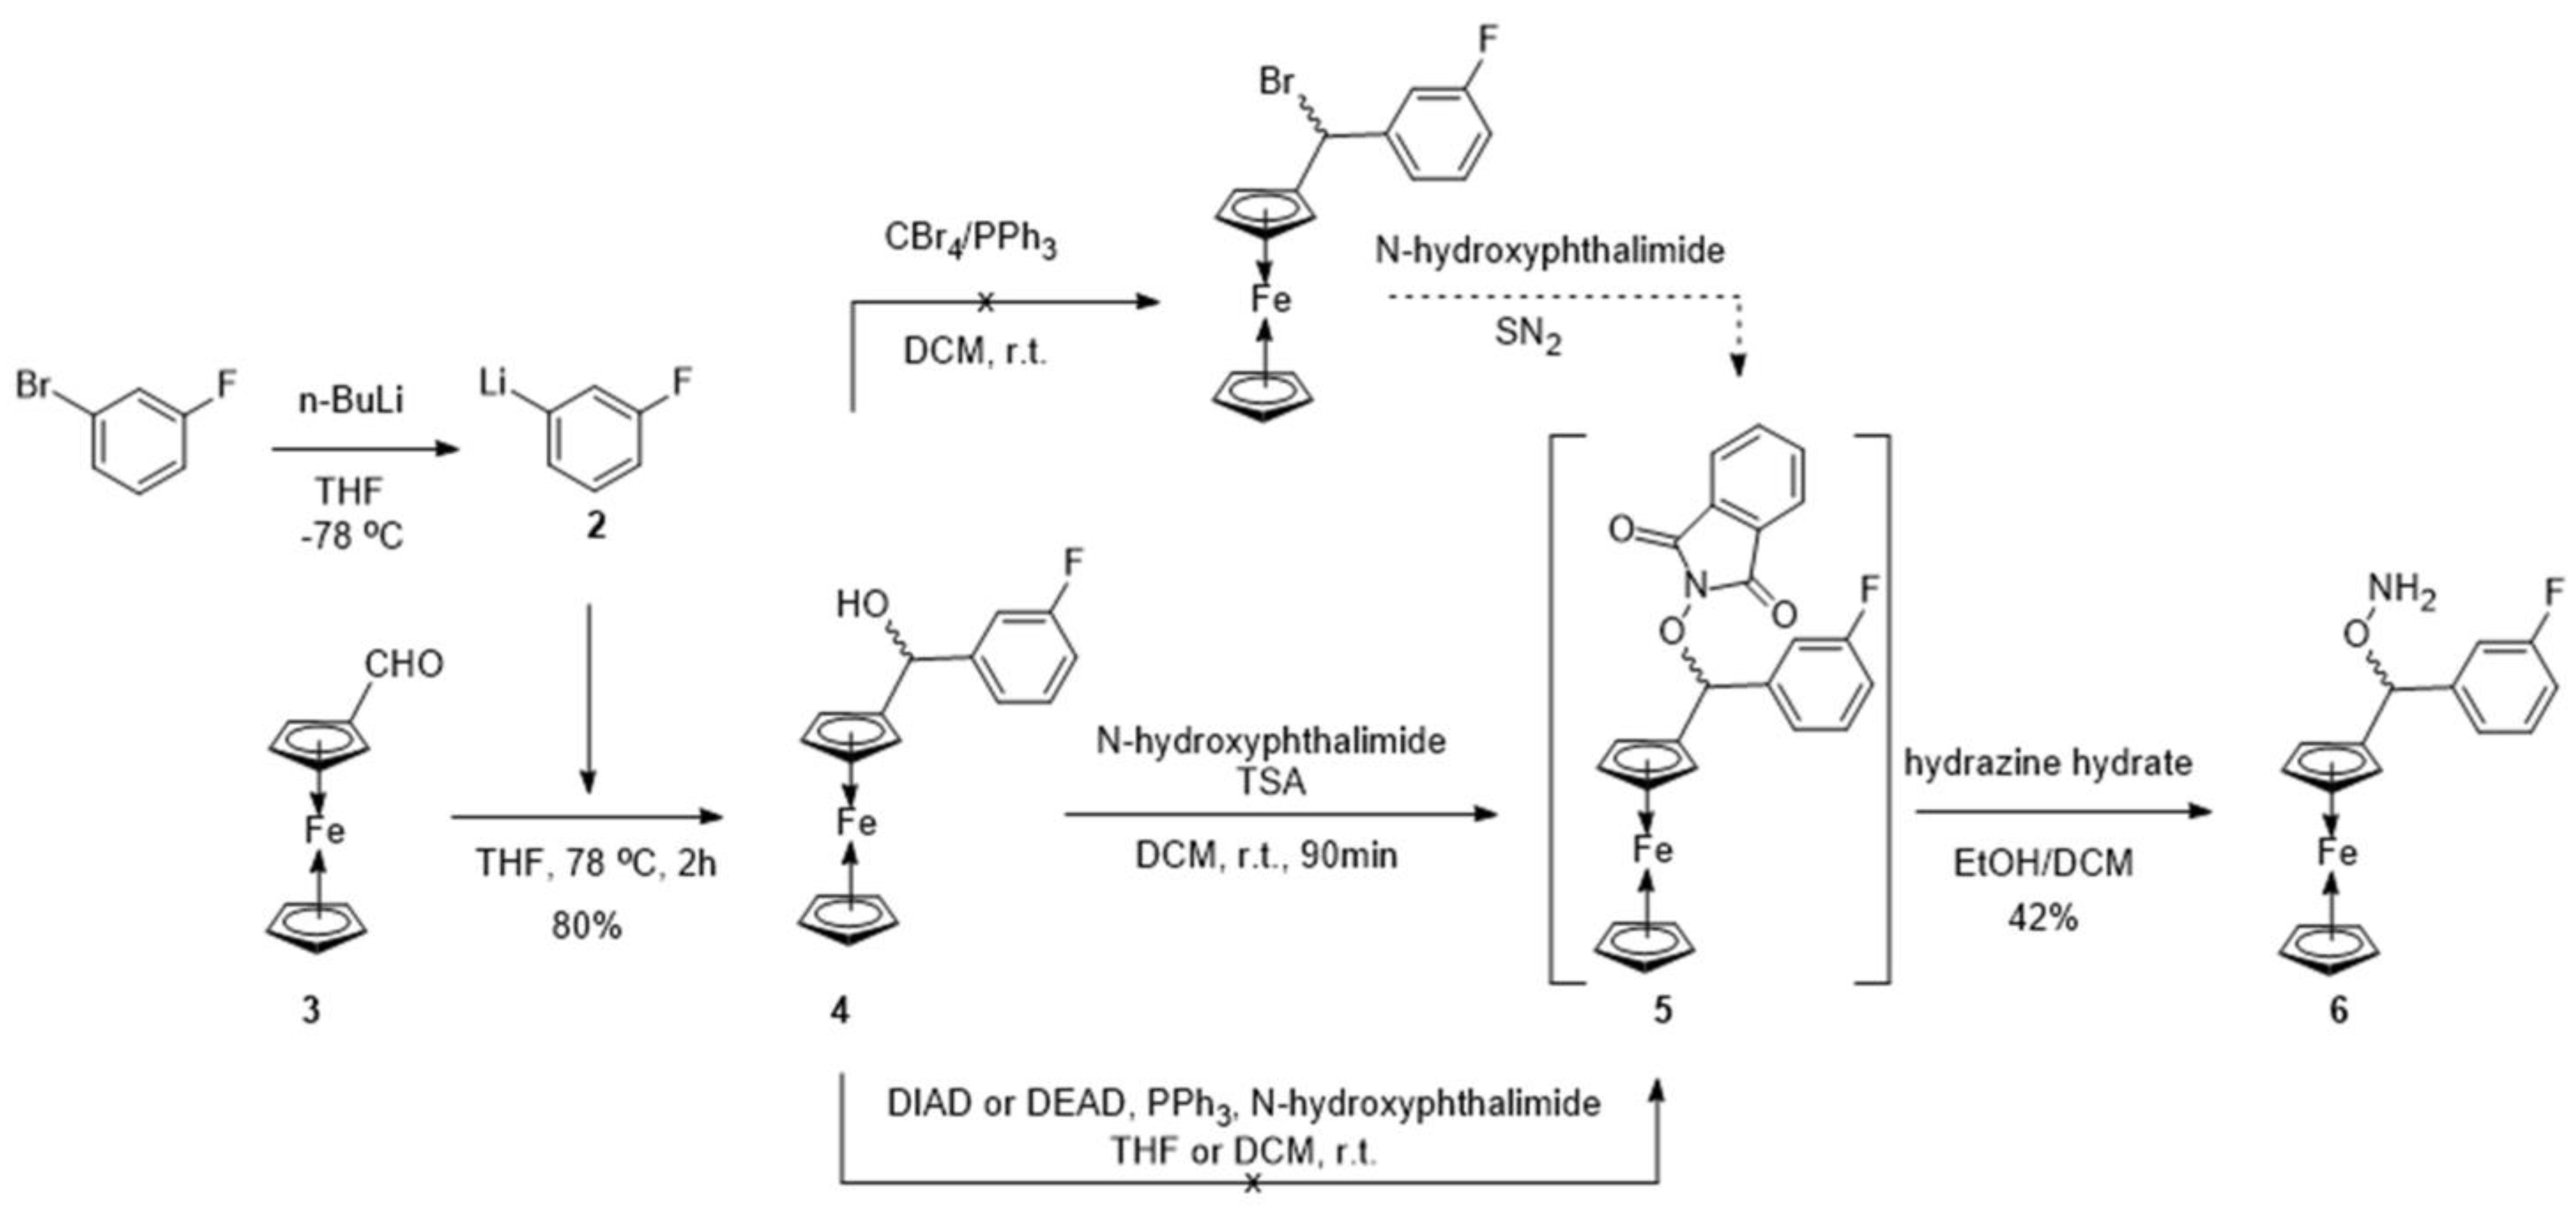
The synthetic route to ferrocene hydroxylamine 6 is shown in Scheme 1 and involves the synthesis of O-alkylated hydroxylamines using N-hydroxyphthalimide following the Gabriel methodology.
Scheme 1.
Synthesis of the ferrocenyl analogue 6.
The reaction of a freshly prepared solution of (3-fluorophenyl)lithium 2 in THF with ferrocenecarboxaldehyde 3 gave alcohol 4 in good yield. Attempts to convert 4 to the corresponding phthalimide 5 under Mitsunobu reaction conditions gave a complex mixture of products, and purification attempts with flash column chromatography or recrystallization did not yield 5, presumably due to its rapid decomposition on silica or from the solvent used for recrystallization. Additionally, efforts to convert 4 to the corresponding ferrocenyl bromide with CBr4/PPh3, with the intention to subsequently react it with N-hydroxyphthalimide through SN reaction to afford 5, also failed to produce the desired bromo compound. The pronounced instability of the phthalyl (5) and bromo intermediates can be attributed to the very strong electron-donating effect of the α-ferrocenyl group, which greatly stabilizes the positive charge on the secondary benzyl cation that favors SN1 substitutions [6,7]. This effect is further exacerbated by the good leaving ability of the bromo and N-hydroxyphthalimide anions.
However, the application of the methodology developed by Reddy et al., which involves the p-TSA catalyzed O-alkylation of hydroxyimides with secondary benzylic alcohols [8], gave a much cleaner reaction profile. Since all efforts to purify 5 in previous attempts failed to give a pure product, the crude reaction mixture was used directly after a workup for the next step, during which it was subjected to hydrazinolysis conditions to provide hydroxylamine 6 in a 42% yield over 2 steps. In contrast to 5, hydroxylamine 6 is stable most probably because the –ONH2 group is a poor leaving group (pKa = 13.7) [9]. Therefore, it can be purified with flash column chromatography, albeit with swift manipulations, and stored.
3. Materials and Methods
3.1. General
All reagents and starting materials were purchased from commercial suppliers and used without further purification. Anhydrous CH2Cl2 was obtained by distillation from calcium hydride under argon. Anhydrous THF was freshly distilled from Na and benzophenone ketyl. All reactions were performed under an inert atmosphere of argon. Concentrated refers to the removal of solvent with a rotary evaporator at normal water aspirator pressure, followed by further evacuation on a high-vacuum line. Thin-layer chromatography was performed using silica gel 60 Å precoated glass-backed plates (0.25 mm thickness) with fluorescent indicators, which were cut. Developed TLC plates were visualized with UV light (254 nm), iodine, or anisaldehyde staining solution. The chromatographic purification of the products was carried out using Fluka silica gel 60 for preparative column chromatography (particle size 40–63 µm). Reactions at 0 °C were carried out in an ice/water bath. Reactions at −78 °C were carried out in a dry ice/acetone bath. Yields refer to spectroscopically pure (>95%) compounds.
NMR spectra were obtained in CDCl3 or DMSO-d6 at 25 °C on a Bruker Avance DRX 500 MHz spectrometer (Supplementary Materials). The measured chemical shifts are reported in δ (ppm), and the residual solvent signal was used as the internal calibration standard (CDCl3): 1H = 7.26 ppm, 13C = 77.18 ppm); (DMSO-d6): 1H = 2.50 ppm. 13C-NMR spectra were measured with complete proton decoupling. Data of NMR spectra were recorded as follows: s = singlet, d = doublet, t = triplet, m = multiplet, dd = doublet of doublet, td = triplet of doublet, and br = broad signal. The coupling constant J is reported in hertz (Hz). 1H- and 13C-NMR peaks were assigned based on the combined analysis of a series of 1H-1H (COSY) and 1H-13C (HSQC, HMBC) correlation spectra.
3.1.1. (Ferrocenyl)(3-fluorophenyl)methanol
n-BuLi (10.2 mL, 16.4 mmol, 1.4 eq., 1.6 M in THF) was added dropwise within 15 min to a solution of 1-bromo-3-fluorobenzene (3.066 g, 17.5 mmol, 1.5 eq) in THF (60 mL) at −78 °C. After the addition was complete, the mixture was stirred for an additional 15 min at the same temperature. Then, a solution of ferrocenecarboxaldehyde (2.5 g, 11.7 mmol, 1 eq.) in THF (30 mL) was added dropwise to the lithiate within 20 min via a cannula. The reaction was allowed to stir for 2 h, during which time the temperature rose to −20 °C. TLC showed complete consumption of the starting material, and the reaction was quenched with saturated NH4Cl solution (10 mL). After reaching room temperature, the mixture was transferred to a separatory funnel with ether (200 mL) and washed with water (2 × 100 mL) and brine (1 × 100 mL). The organic layer was dried under Na2SO4, filtered, and concentrated to dryness. The brown oily residue was purified by flash column chromatography using hexanes/AcOEt 9:1, increasing to 8:2 to afford the title compound (2.9 g, 80%) as a deep-orange solid. m.p.: 63–65 °C; 1H-NMR (CDCl3), δ(ppm): 7.25–7.30 (m, 1H, 5-Har), 7.11–7.15 (m, 2H, 2,6-Har), 6.93 (td, J = 8.4, 2.2 Hz, 1H, 4-Har), 5.39 (s, 1H, ArCH(OH)), 4.23–4.29 (m, 9H, C5H4, C5H5), 2.42 (br, 1H, OH); 13C-NMR (CDCl3), δ(ppm): 162.92 (d, 1JC-F = 244.5 Hz, 3-Car), 146.0 (d, 3JC-F = 7.1 Hz, 1-Car), 129.79 (d, 3JC-F = 7.9 Hz, 5-Car), 121.94 (6-Car), 114.34 (d, 2JC-F = 22.2 Hz, 4-Car), 113.23 (d, 2JC-F = 22.2 Hz, 2-Car), 94.28 (ArCH(OH)), 71.46 (C5H4), 68.84 (C5H5), 68.58 (2 × C5H4), 67.81 (C5H4), 66.02 (C5H4). HRMS: m/z calculated for C17H14FFe [M + H − H2O]+: 293.0429, found: 293.0421.
3.1.2. 2-((Ferrocenyl)(3-fluorophenyl)methoxy)isoindoline-1,3-dione
To a solution of alcohol 4 (0.5 g, 1.61 mmol, 1 eq.) in DCM (8 mL), N-hydroxyphthalimide (394 mg, 2.42 mmol, 1.5 eq.) was added. After stirring for 10 min, p-TSA (30.6 mg, 0.16 mmol, 0.1 eq) was added to the mixture, and the reaction was stirred for 90 min, at which point the reaction’s color was almost black. The mixture was diluted with ether (60 mL) and washed with sat. NaHCO3 solution (3 × 30 mL) and brine (1 × 30 mL), dried over Na2SO4, filtered, and concentrated to dryness. The orange residue (0.7 g) was used directly for the hydrazinolysis step.
3.1.3. O-((Ferrocenyl)(3-fluorophenyl)methyl)hydroxylamine
The crude material from the previous step (0.7 g) was dissolved in ethanol (2 mL) and DCM (2 mL), and aq. hydrazine monohydrate (55% w/v, 0.37 mL, 4.02 mmol) was added. After 30 min of stirring, a white precipitate appeared while stirring was continued for an additional 90 min, at which point TLC confirmed the complete consumption of the starting material. Subsequently, ether (30 mL) was added, and the reaction mixture was placed in the freezer for 5 h. The solid was then removed by suction filtration, and the filtrate was concentrated to dryness. The crude material was purified by flash column chromatography by loading it neat on the column and using hexane/AcOEt/Et3N 8:2:0.1 to provide the title compound (0.22 g, 42% over two steps) as an orange solid. m.p.: 87–88 °C; 1H-NMR (DMSO-d6), δ(ppm): 7.40 (dd, J = 7.8, 6.4 Hz, 1H, 5-Har), 7.23 (d, J = 7.6 Hz, 1H, 6-Har), 7.19 (d, J = 9.9 Hz, 1H, 2-Har), 7.10 (td, J = 8.5, 1.9 Hz, 1H, 4-Har), 6.03 (br, 2H, ONH2), 5.30 (s, 1H, ArCH(ONH2), 4.19 (s, C5H4), 4.09–4.13 (m, 7H, C5H4, C5H5), 3.94 (s 1H, C5H4); 13C-NMR (CDCl3), δ(ppm): 163.0 (d, 1JC-F = 246.2 Hz, 3-Car), 144.04 (d, 3JC-F = 6.3 Hz, 1-Car), 129.91 (d, 3JC-F = 7.7 Hz, 5-Car), 123.15 (6-Car), 114.76 (d, 2JC-F = 21.2 Hz, 4-Car), 114.13 (d, 2JC-F = 21.7 Hz, 2-Car), 87.36 (C5H4), 85.17 (ArCH(ONH2), 68.91 (C5H5), 68.40 (C5H4), 68.30 (C5H4), 68.27 (C5H4), 67.39 (C5H4). HRMS: m/z calculated for C17H16FFeNO [M]+: 325.0565, found: 325.0561.
4. Conclusions
O-((ferrocenyl)(3-fluorophenyl)methyl)hydroxylamine, a ferrocenyl analogue of the diaryl hydroxylamine scaffold was synthesized as an attempt to enhance the pan-inhibitory potency of the diaryl hydroxylamine scaffold. The methodology described herein enables the synthesis of O-((ferrocenyl)(aryl)methyl)hydroxylamines whose synthesis has not yet been reported in the literature. The compound is currently under biological evaluation.
Supplementary Materials
The following material is available online. Figure S1: 1H-NMR spectrum of compound 4; Figure S2: 13C-NMR spectrum of compound 4; Figure S3: HRMS spectrum of compound 4; Figure S4: 1H-NMR spectrum of compound 6; Figure S5: 13C-NMR spectrum of compound 6; Figure S6: HRMS of compound 6.
Author Contributions
Design, conception, and writing, A.C. and M.S.P.; synthesis, structure elucidation, and NMR studies, A.S.F., M.G. and A.C. All authors have read and agreed to the published version of the manuscript.
Funding
This research is co-financed by Greece and the European Union (European Social Fund- ESF) through the Operational Programme “Human Resources Development, Education and Lifelong Learning 2014-2020” in the context of the project “Targeting the kynurenine pathway for tumor imaging and characterization by single-photon emission computed tomography (SPECT) and positron emission tomography (PET)” (MIS 5047830).
Conflicts of Interest
The authors declare no conflict of interest.
References
- Singh, A.; Lumb, I.; Mehra, V.; Kumar, V. Ferrocene-appended pharmacophores: An exciting approach for modulating the biological potential of organic scaffolds. Dalton Trans. 2019, 48, 2840–2860. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Kilpin, K.J.; Dyson, P.J. Enzyme inhibition by metal complexes: Concepts, strategies and applications. Chem. Sci. 2013, 4, 1410–1419. [Google Scholar] [CrossRef] [Scilit]
- Patra, M.; Gasser, G. The medicinal chemistry of ferrocene and its derivatives. Nat. Rev. Chem. 2017, 1, 0066. [Google Scholar] [CrossRef] [Scilit]
- Lewis-Ballester, A.; Pham, K.N.; Batabyal, D.; Karkashon, S.; Bonanno, J.B.; Poulos, T.L.; Yeh, S.R. Structural insights into substrate and inhibitor binding sites in human indoleamine 2,3-dioxygenase. 1. Nat. Commun. 2017, 8, 1693. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Winters, M.; DuHadaway, J.B.; Pham, K.N.; Lewis-Ballester, A.; Badir, S.; Wai, J.; Sheikh, E.; Yeh, S.R.; Prendergast, G.C.; Muller, A.J.; et al. Diaryl hydroxylamines as pan or dual inhibitors of indoleamine 2,3-dioxygenase-1, indoleamine 2,3-dioxygenase-2 and tryptophan dioxygenase. Eur. J. Med. Chem. 2019, 162, 455–464. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Juric, S.; Marijan, M.; Kronja, O. Electrofugality of Some Ferrocenylphenylmethyl Cations. Croat. Chem. Acta 2019, 92, 307–313. [Google Scholar] [CrossRef] [Scilit]
- Marijan, M.; Juric, S.; Mihalic, Z.; Kronja, O. Impact of the alpha-Ferrocenyl Group on the Solvolytic Reactivity-Electrofugality-of Ferrocenylphenylmethyl Cations. Eur. J. Org. Chem. 2019, 2019, 537–546. [Google Scholar] [CrossRef] [Scilit]
- Reddy, C.R.; Radhika, L.; Kumar, T.P.; Chandrasekhar, S. First Acid-Catalyzed Entry to O-Alkylated Hydroximides from Benzylic Alcohols. Eur. J. Org. Chem. 2011, 2011, 5967–5970. [Google Scholar] [CrossRef] [Scilit]
- Lind, J.; Merenyi, G. Kinetic and thermodynamic properties of the aminoxyl (NH2O) radical. J. Phys. Chem. A 2006, 110, 192–197. [Google Scholar] [CrossRef] [Scilit]
Publisher’s Note: MDPI stays neutral with regard to jurisdictional claims in published maps and institutional affiliations. |
© 2022 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license (https://creativecommons.org/licenses/by/4.0/).