


Abstract
While either a spin or point-group adaptation is straightforward when considered independently, the standard technique for factoring isotropic spin Hamiltonians by the total spin S and the irreducible representation of the point group is limited by the complexity of the transformations between different coupling schemes that are related in terms of their site permutations. To overcome these challenges, we apply projection operators directly to uncoupled basis states, enabling the simultaneous treatment of spin and point-group symmetry without the need for recoupling transformations. This provides a simple and efficient approach for the exact diagonalization of isotropic spin models, which we illustrate, with applications in Heisenberg spin rings and polyhedra, including systems that are computationally inaccessible with conventional coupling techniques.
1. Introduction
With the emergence of the field of molecular magnetism [1,2], the rapid expansion in the synthesis of exchange-coupled clusters, along with their physical and spectroscopic characterization—such as their temperature-dependent magnetic susceptibilities, electron paramagnetic resonance (EPR), and inelastic neutron-scattering spectra—has made theoretical modeling based on effectively treating the open-shell metal ions as spin centers increasingly important [3,4]. The most common description is the Heisenberg model of pairwise scalar coupling, Equation (1),
where is a local spin vector, which also arises in other contexts, such as the coupling between nuclear spins in NMR [5]. While the direction-dependent properties observed in EPR require the inclusion of anisotropic terms, these effects can often be treated via perturbation theory [3].
The levels of such an isotropic Hamiltonian are characterized by a total-spin value S and comprise states with magnetic quantum numbers (eigenvalues of the z-component ). Molecular symmetry manifests in terms of the permutations of spin sites [6,7], allowing each level to be additionally assigned to an irreducible representation (irrep) of the respective point group (PG). Point-group symmetry follows from choosing the isotropic couplings in a way that reflects the desired molecular symmetry; anisotropic interactions explicitly encode the geometric structure [8,9].
Given the rapidly increasing dimensions of the Hilbert space with the number of centers, it becomes essential, when aiming for exact diagonalization (ED), to exploit all available symmetries to factor the Hamiltonian into smaller blocks, each characterized by labels S and [10]. Beyond facilitating computations, a classification of levels can be useful for deriving selection rules [11,12] or for separating the geometric and dynamic features of spectra [11,12,13].
Combining PG and symmetry by working from a basis of uncoupled configurations (also known as a Zeeman product basis [5]) defined by local z-projections with the sum M, as in Equation (2),
is straightforward [6,14]. In addition, for , spin-flip symmetry can be used to separate sectors with an even or odd S [14]. Such approaches have been applied widely, e.g., in the full ED of symmetric polyhedra [15] or in calculations of the thermodynamic properties of lattice fragments with up to sites using the finite-temperature Lanczos method (FTLM) [16]. However, factorization of the Hamiltonian in terms of and S, which involves constructing eigenfunctions of (rather than only ), is considered more demanding and has been pursued in just a few cases. The most prominent strategy used for this task [6,10,17,18,19] is based on genealogical coupling (GC) [3,20], where individual spins are coupled into subunits of increasing size until a multiplet with the total spin S is obtained. The calculation of the Hamiltonian elements from such a basis makes use of irreducible tensor–operator techniques and proceeds by successive decoupling through Wigner-9j symbols [3]. However, a complication arises from PG adaption [6]: for most systems, there is no compatible coupling scheme, i.e., PG operations may produce states belonging to different schemes, related to the original basis by their respective site permutations, and transforming them back can become prohibitively demanding as it requires the computation of a large number of square roots and Wigner-6j symbols [10,17]. Resorting to the use of a subgroup that enables the construction of a compatible coupling scheme [6,10] to avoid such transformations presents two drawbacks: it neither produces the smallest possible Hamiltonian matrix blocks, nor does it provide a comprehensive labeling of eigenstates.
GC is not the only method used for constructing spin eigenfunctions [20], but Sahoo et al. [21] briefly reviewed why various other techniques are also difficult to apply in conjunction with arbitrary point groups. To overcome this limitation, these authors suggested transformations between uncoupled and valence bond (VB) states. In a VB state, each site is replaced by auxiliary spin-1/2 objects, of which are singlet-paired according to Rumer–Pauling rules, supplemented by the restriction that no pairs are formed within the same center. As PG symmetrization and setting up the Hamiltonian would require complicated transformations between different diagrams, VB states are transformed into the uncoupled basis set (using Clebsch–Gordan coefficients), where the PG projectors and the Hamiltonian have simple representations. Thus, coupled and uncoupled bases are utilized for spin and PG adaptation, respectively. This approach is extendable to fermionic systems [22] and can be applied to all types of point groups, but it has not been widely adopted. Since full ED was only demonstrated for a rather small system with a cubic arrangement of 14 sites (the dimension of the largest symmetry subspace was 219) [21], it remains uncertain how well the procedure would scale to more challenging cases.
Our present strategy of applying spin and PG projectors to uncoupled states is suitable for arbitrary point groups, and it is simpler than the GC or VB methods. Note that Bernu et al. [23] combined Löwdin’s spin-projection operator [24]— in Equation (6) below—with a spatial-symmetry projector in a Lanczos process to compute a small number of lowest states for clusters of up to 36 spin-1/2 centers. We recently used a similar approach to obtain analytical solutions for particularly small clusters [25]. In contrast, the present aim is the full numerical diagonalization of systems that would be challenging to solve without using all available symmetries. Details on forming spin eigenfunctions based on a group-theoretical formulation of (a procedure that we investigated recently [26]) and on the construction of the Hamiltonian in each subspace are provided in the following Theory and Computational Details section. In the Results and Discussion section, we assess the numerical accuracy of this approach and present several cases that represent some of the largest fully solved spin models to date, including systems that cannot be practically handled using the conventional GC method.
2. Theory and Computational Details
In the following, we explain the combined spin and PG adaptation of uncoupled configurations to set up a generalized eigenvalue problem (GEP) defined by the Hamiltonian and the overlap matrix in a symmetry subspace characterized by the quantum numbers S and . We start with a discussion of PG adaptation. The projection onto a specific component of a possibly multidimensional irrep is given in Equation (3):
where h is the order of the group, the sum runs over all elements g, is a symmetry operation that permutes the z-projections , and is a diagonal element of the representation matrix (the asterisk * denotes complex conjugation). Note that, when employing characters instead of specific matrix elements , the components are not separated; see Equation (4).
We work with Equation (3) instead of Equation (4) in order to reduce the size of the Hamiltonian matrix in each respective sector by the number of components n (the dimension of ), i.e., . The subspace dimension can be represented as the weighted sum of Equation (5), where is the matrix representation of in the basis set with the selected M. To compute the traces of individual operations, , we examine each and check whether it is mapped onto itself under the action of .
The irreps of the cyclic group , which is relevant for spin rings, are one-dimensional and characterized by a crystal momentum associated with the eigenvalues of the site permutation corresponding to a rotation of . Except when (Mulliken label A) or (label B, for even N), k and comprise degenerate pairs spanning two-dimensional irreps (, , etc.) of the dihedral group . Therefore, when working with the group, ED can be restricted to the sectors for even N or for odd N. A few guidelines for the simple construction of the required matrices for more complicated groups like icosahedral or octahedral were provided in our previous work [26]: permutation and representation matrices [27] are explicitly set up only for the group generators, and the permutations are pairwise-compounded until a closed set of elements is obtained; whenever a new element is found, the corresponding pair of representation matrices is similarly multiplied to obtain the new representation matrix.
In many cases—including in cyclic groups—a complete and orthogonal basis in a sector can be constructed by ensuring that each uncoupled state appears in at most one linear combination. The configurations selected by this rule form a complete orthogonal space upon the application of and are stored in the columns of the matrix (each column of has exactly one entry of 1). While it was previously assumed that each would always contribute to at most one linear combination [25], we have observed that this does not generally hold for multidimensional irreps of the icosahedral group, where a may appear in multiple states for a given , provided its coefficients have different magnitudes in different states. A small threshold on the number of these distinct absolute amplitudes (e.g., two or three) suffices to generate the complete space. The resulting is generally overcomplete with respect to . However, rather than directly pruning linear dependencies, a single rank-revealing selection (as discussed below) is performed later to choose configurations for simultaneous spin and PG projection.
For large , we typically stop the generation of once approximately 20% more states have been collected than the number of levels. If the subsequent spin adaptation step fails to fully span the space with this truncated set, then additional configurations can be considered. However, we found that to produce a suitable truncated set, the should be processed in a random order, not sequentially. (For example, for , a listing according to a binary representation, where and represent 1 and 0, would not be adequate; the ordering should be randomized for generating a truncated PG-adapted basis.)
Turning now to the construction of spin eigenfunctions, we note that the direct application of Löwdin’s projector, Equation (6),
can induce significant numerical rounding errors due to the repeated matrix–vector multiplications needed to isolate the subspace with the desired S [23,26]. To circumvent these issues, we employ a group-theoretical integral formulation of [28],
where is a diagonal element of the Wigner rotation matrix for spin S. Evaluating the wave function , i.e., Equation (8), obtained by applying to a state with , is greatly simplified by the integral formulation.
The integrals over Euler angles and (corresponding to rotations about the z-axis) are trivial. The remaining integration over involves the element of the small Wigner matrix and is reduced to a standard integral for spin-1/2 systems [29], which leads to a closed-form expression for , Equation (9), in terms of a special kind of so-called Sanibel coefficient [30] involving only a sign factor and a binomial coefficient,
where k is the number of sites where and is the number of sites in the reference configuration, i.e., , . For , the wave function becomes a linear combination of standard integrals [26] (one may alternatively evaluate the integrals numerically on a grid). As we recently demonstrated [26], this analytical group-theoretical approach to evaluating the action of provides numerical advantages over Löwdin’s projector. Accordingly, we exclusively employ this method in the present work.
The number of multiplets with spin S transforming according to is calculated from Equation (5), assuming . The level is symmetric under all spin-permutations; thus, , where is the totally symmetric irrep.
A subset (see the next section on the selection procedure) of columns from comprises the matrix R. These configurations span a complete, linearly independent basis once the projectors have been applied. With and denoting their respective matrices (here and in the following, to avoid clutter, we will typically just refer to Γ for simplicity and let denote , while keeping in mind that the projection is always performed onto a specific component μ), and considering the Hermiticity and idempotency of projection operators, the Hamiltonian and overlap matrices in a sector are formulated as in Equations (11) and (12).
The Hamiltonian H (in the uncoupled basis where ) is generated in a sparse format and is multiplied by the PG adapted states , which are also in a sparse format. The spin adaptation works as described above, cf. Equation (9) for , but as spin eigenfunctions are dense vectors, fully storing may require excessive memory. Therefore, a single row (or a limited number of rows) of is multiplied with the entire to construct (as well as ) sequentially. A symmetrization of and should be performed to eliminate numerical rounding errors, which would slow down the subsequent solution of the GEP and could lead to small imaginary components in the eigenvalues. The nonorthogonality of the symmetry-adapted basis is a feature shared with the GC and VB methods mentioned in the Introduction [17,21]. Rather than performing an explicit orthogonalization, we solve the GEP using the eig function in MATLAB R2024a.
We now discuss the selection of states from for the combined application of and . As explained, the columns of span the (possibly truncated) space under . Note that for pure spin projection in spin-1/2 systems, the so-called Löwdin theorem specifies how configurations ought to be selected so that under the action of they span a complete, linearly independent basis in the sector [20]. We have recently generalized Löwdin’s theorem to systems with arbitrary local spin quantum numbers [26] but observed that it does not constitute the most numerically stable procedure in terms of the conditioning of the overlap matrix (i.e., eigenvalues can approach zero within numerical precision). Instead, we proposed an iterative pivoted Cholesky decomposition [31,32] (PCD). This strategy efficiently selects a linearly independent subset for constructing a well-conditioned GEP. A practical alternative is a rank-revealing QR factorization with column-pivoting, which is directly available in MATLAB without additional coding, unlike PCD.
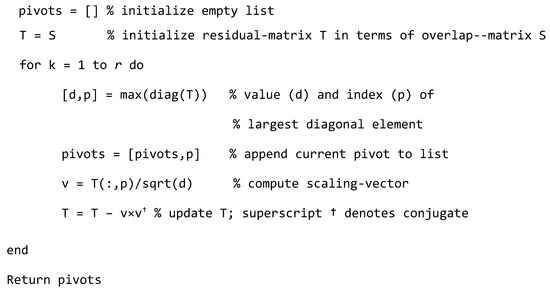
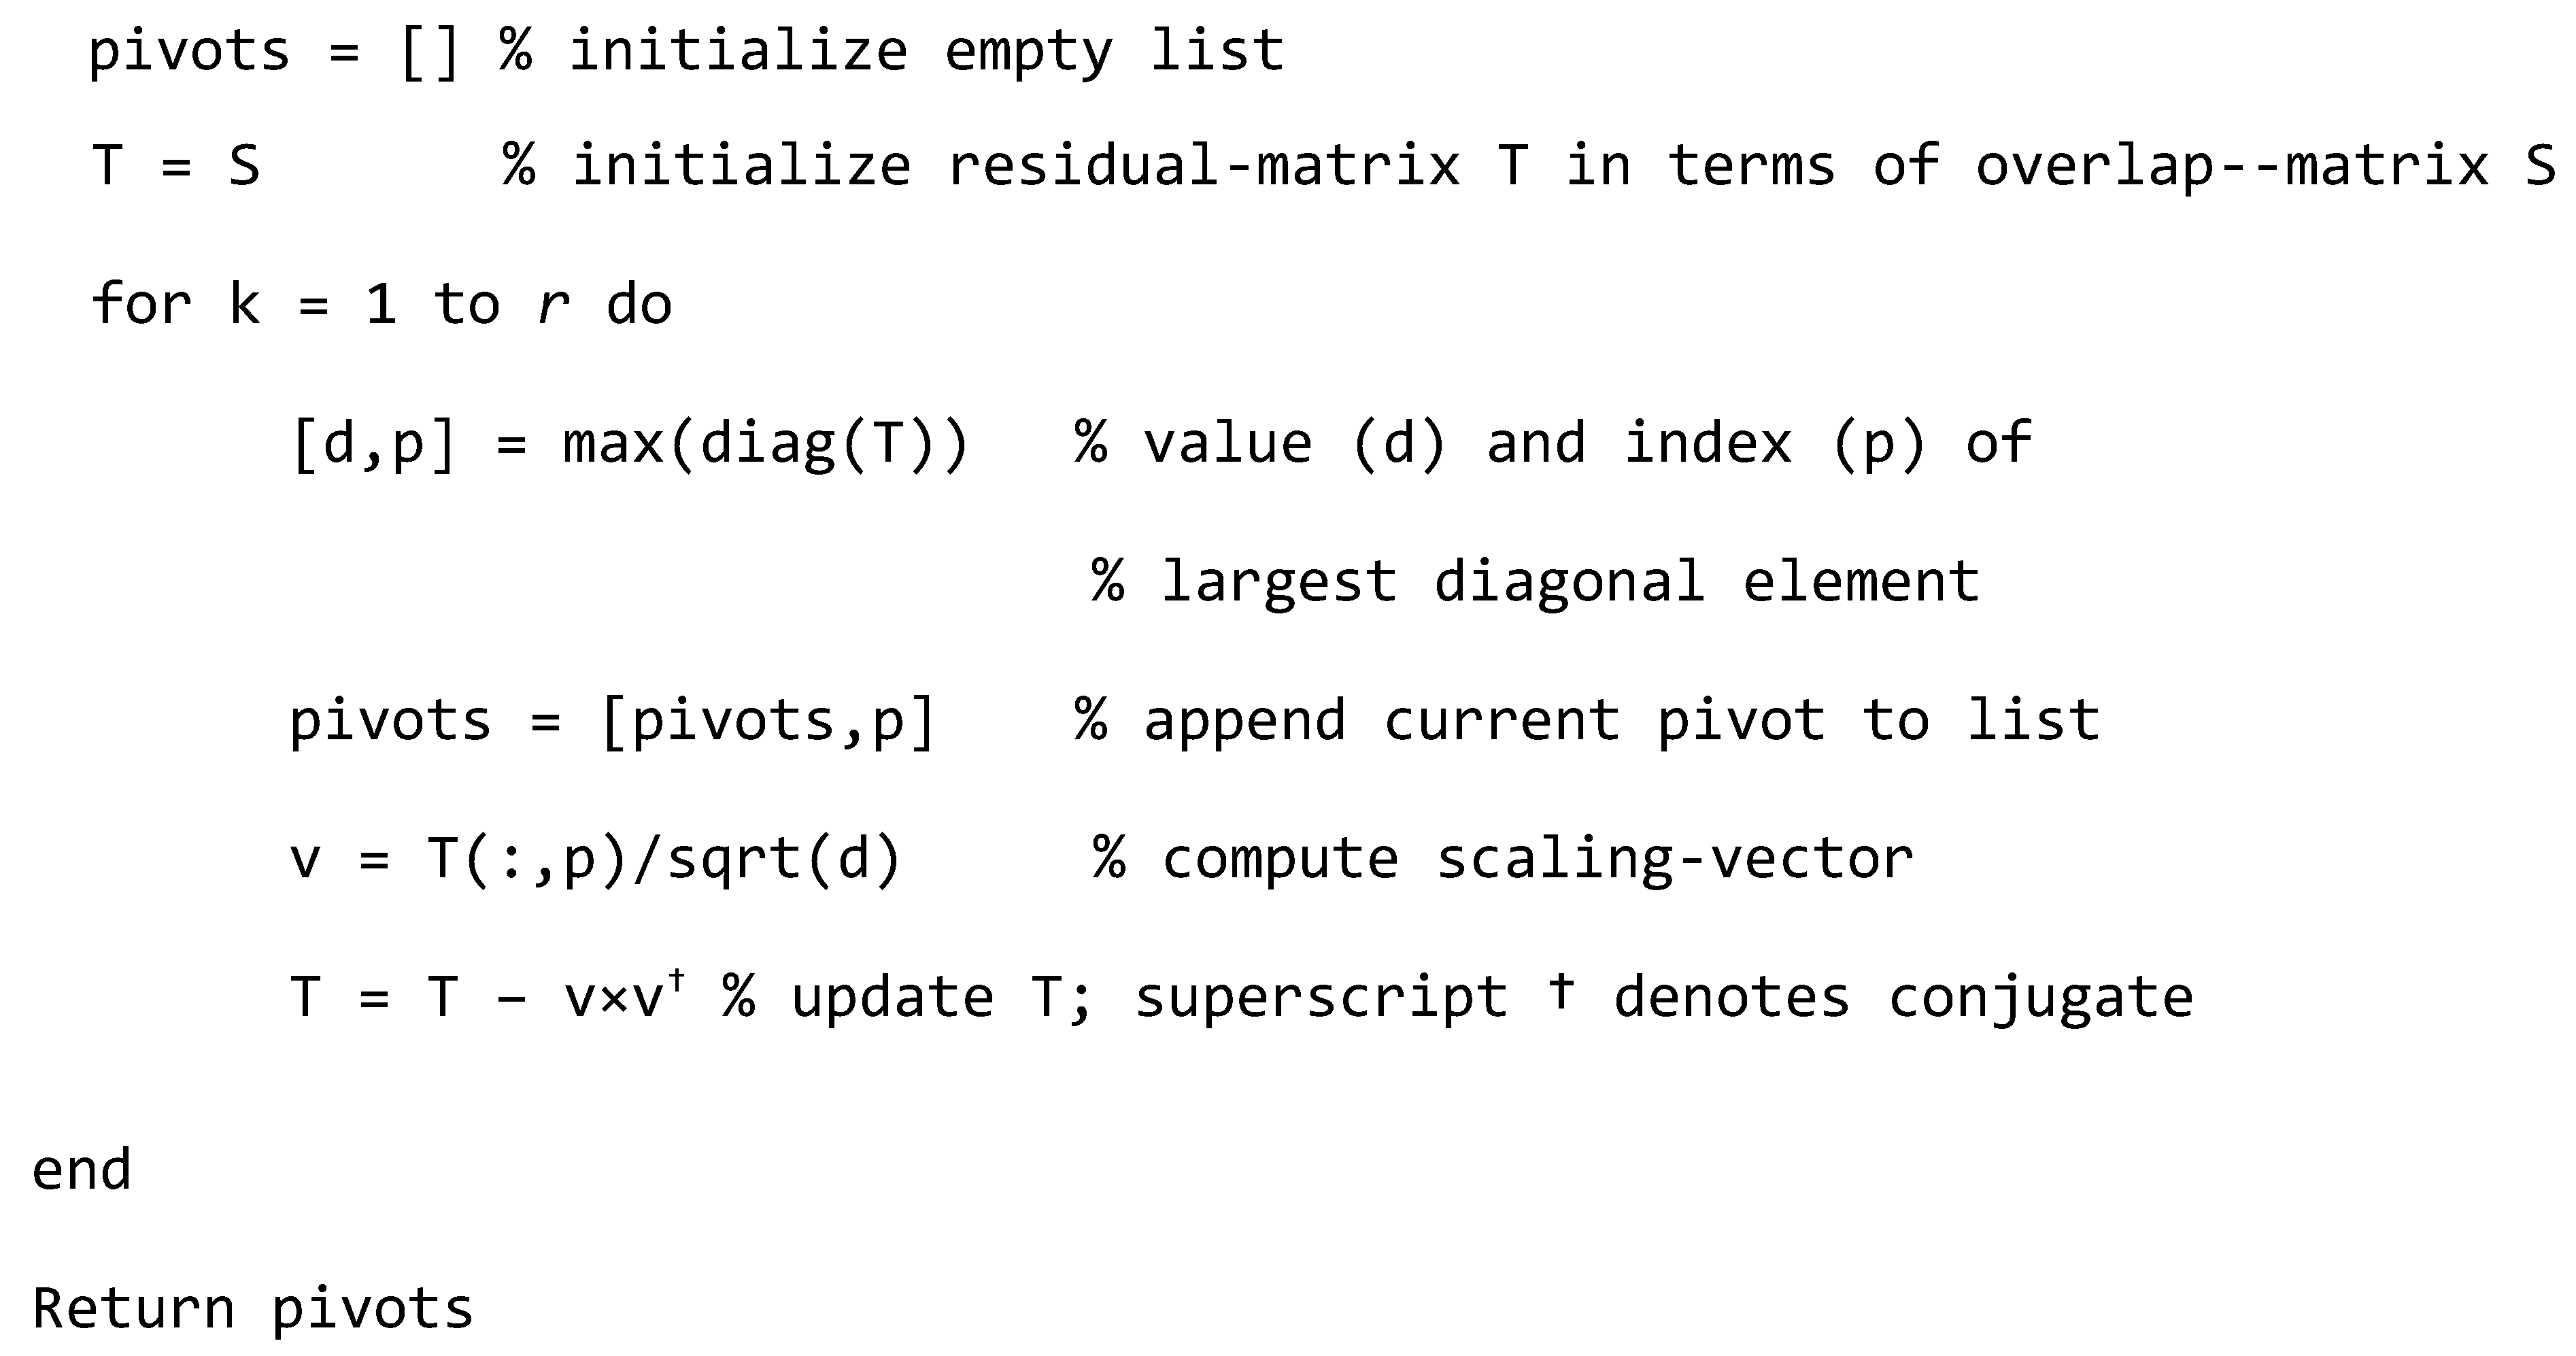
To our knowledge, there is no analogue to Löwdin’s theorem for selecting configurations for the combined action of and , and we thus build on the iterative strategy described previously to construct a basis [26]. However, this procedure is now applied in a slightly modified fashion (our explanation focuses specifically on PCD, but the iterative approach for state selection could be performed similarly for QR factorization with column-pivoting): rather than initially selecting a minimal number of candidate states from , we choose roughly 10–20% more to reduce the likelihood of multiple iterations (if is a complete set of PG-adapted states generated from an ordered list of uncoupled configurations, its columns should be selected randomly). From these selected states, provisional overlap and Hamiltonian matrices and are formed, and the rank r of is computed at a reasonable tolerance well above numerical accuracy. At each step, the PCD routine identifies the pivot—the index of the largest diagonal element of the current residual matrix (initially set to )—and then forms a normalized scaling vector to update that matrix. This procedure is repeated until r pivots have been recorded, as detailed in Figure 1. States failing the selection are removed, and if the rank of in the subset of the selected pivot indices remains below , then an additional batch of states is appended. If the process stagnates, the tolerance for rank determination may be reduced. Each iteration only updates and for any newly added states, thereby avoiding a recomputation of matrix elements.
Figure 1.
MATLAB-like pseudocode for a pivoted Cholesky decomposition (PCD). Starting from the overlap matrix S, the algorithm iteratively selects r basis states to span a linearly independent space by updating the residual matrix T and tracking the pivot indices.
In our implementation, the construction of spin eigenstates by analytically evaluating the group-theoretical integral of Equation (7) tends to lead to a computational bottleneck for . We therefore accelerate the construction of and by processing uncoupled states in parallel. With respect to assembling and solving the GEP, different sectors are independent—aside from the minor overhead of constructing the same uncoupled basis and Hamiltonian H for a given S but different Γ—thus permitting the straightforward distribution of these computations across the nodes of a computer cluster.
3. Results and Discussion
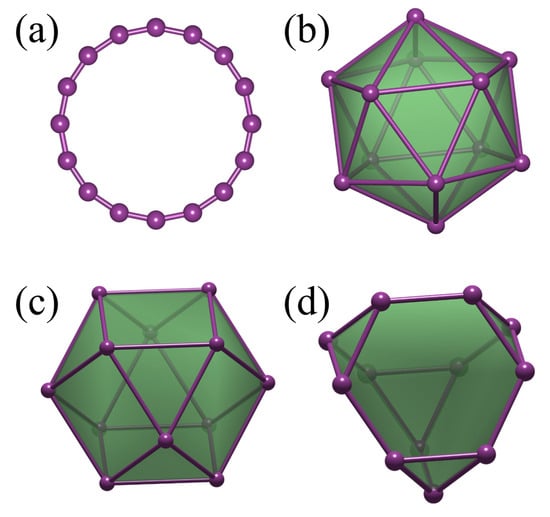
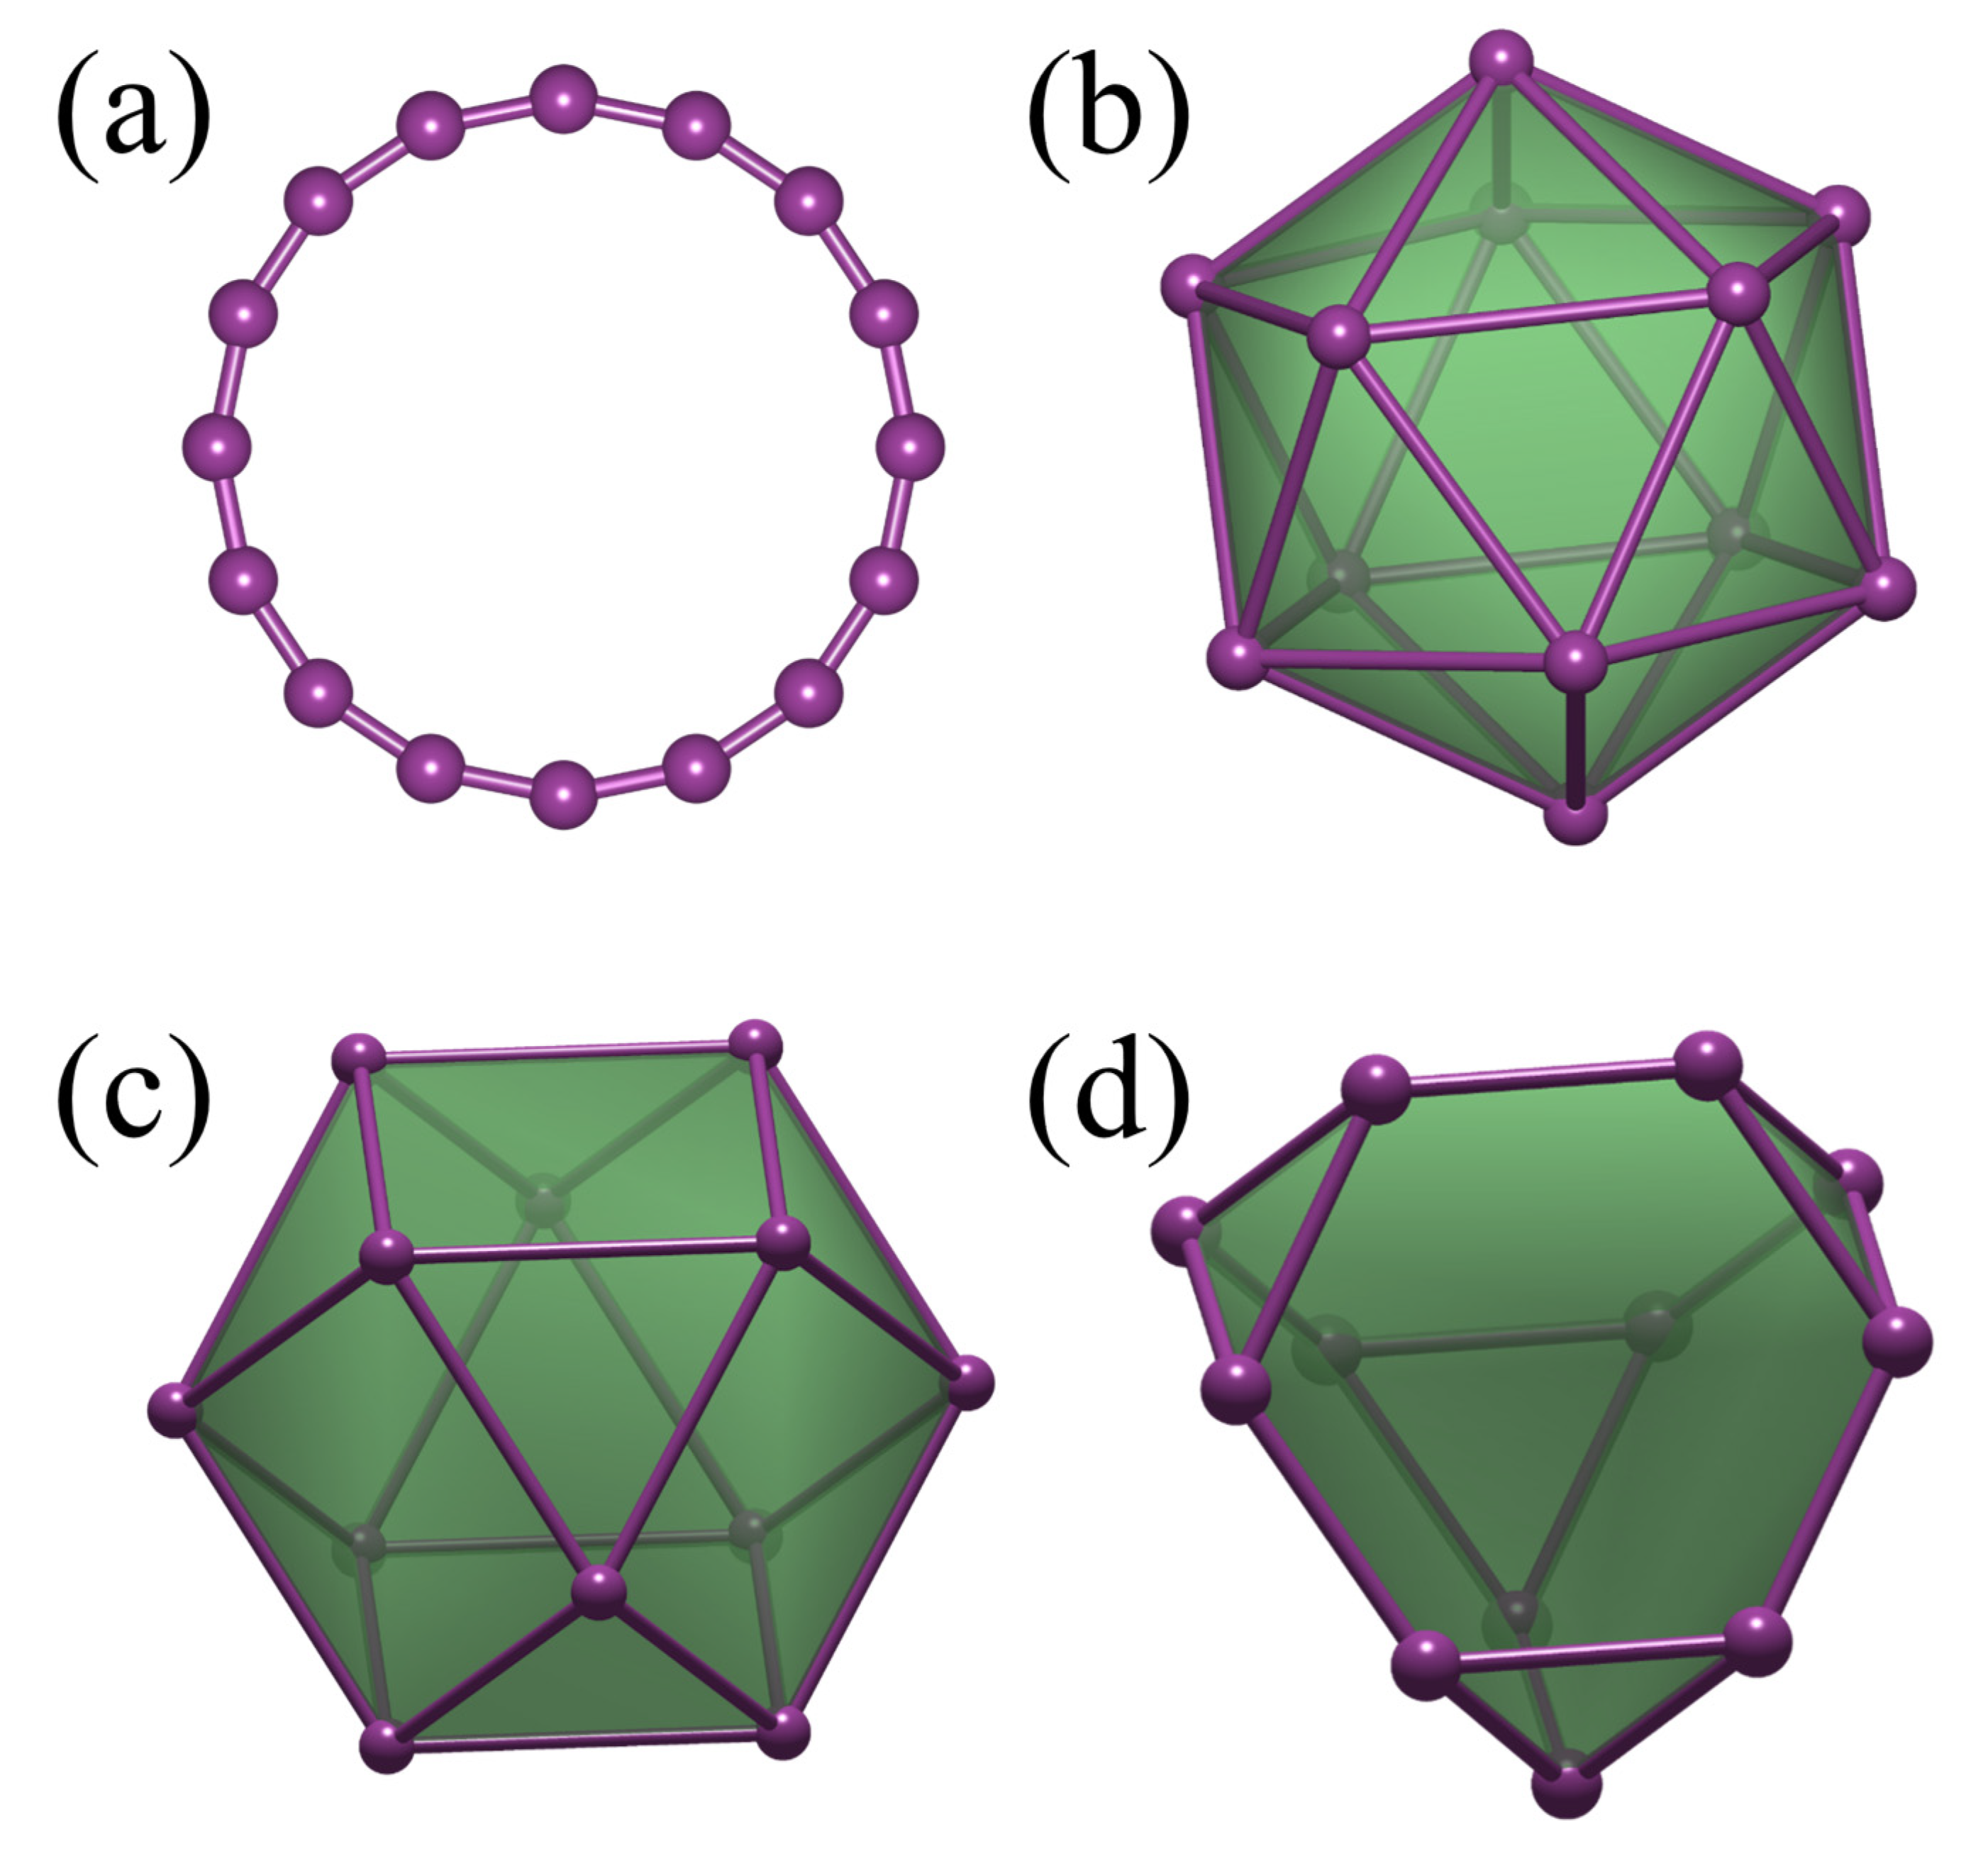
We consider spin rings and polyhedra—specifically, the icosahedron, cuboctahedron and truncated tetrahedron (see Figure 2)—to illustrate the exact diagonalization procedure that uses a complete symmetry factorization of the Heisenberg spin Hamiltonian. Throughout, we assume nearest-neighbor couplings with , which represent antiferromagnetic interactions. We note that our projection operator approach applies to any spin Hamiltonian exhibiting both total spin and point-group symmetries and is therefore not limited to the antiferromagnetic Heisenberg model.
Figure 2.
Heisenberg spin clusters considered in this work: (a) ring, (b) icosahedron, (c) cuboctahedron, (d) truncated tetrahedron. Spheres denote spin sites, and lines represent antiferromagnetic couplings with .
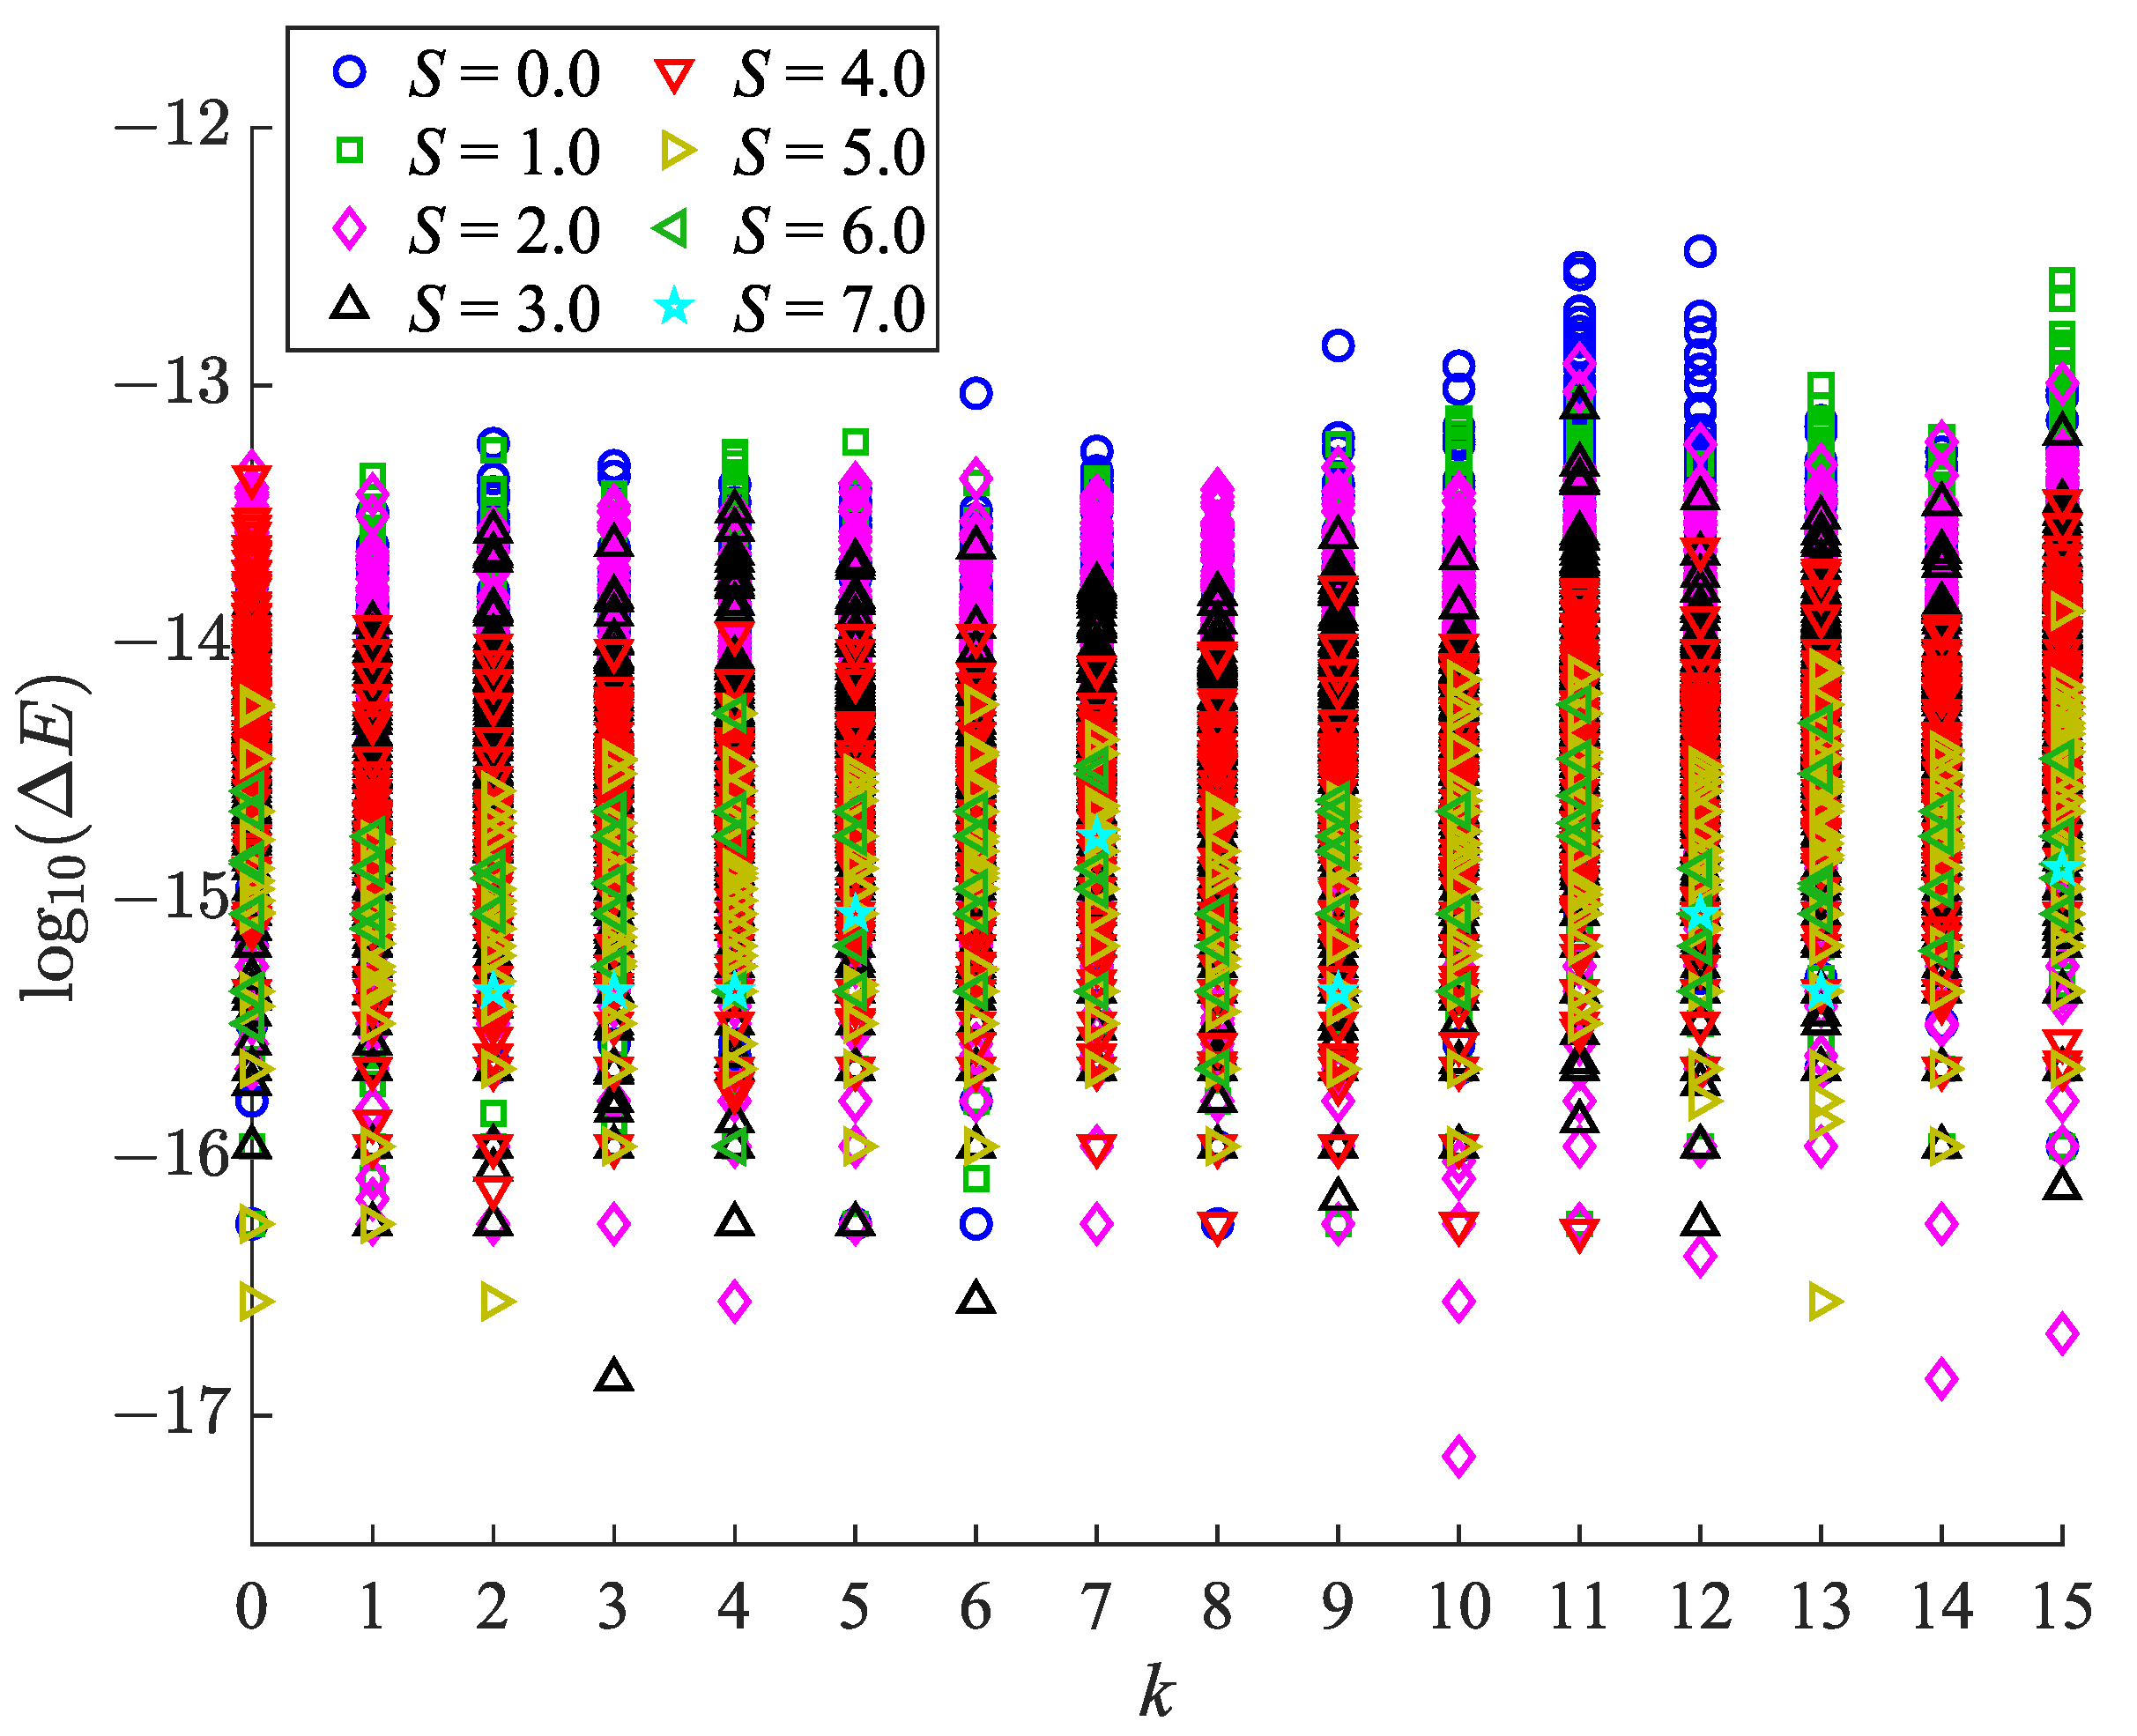
Spin ring. As the first example, we compare the results of two methods when factoring the spin Hamiltonian for an , ring: (1) using the straightforward PG adaptation of uncoupled states, labeling sectors by (quantum numbers S can be assigned to the levels using one of two methods: (i) calculating the spectra in all subspaces, and if a level appears in a specific M subspace but not in , it corresponds to , or (ii) diagonalizing only in the space with the smallest absolute value of M ( or ) and assigning S from the expectation value , which requires access to the eigenfunctions); and (2) combining PG adaptation with total spin projection, labeling sectors by . The first method involves a standard eigenvalue problem, while the second requires solving a generalized eigenvalue problem (GEP). The numerical accuracy of the approach is guaranteed to be at approximately machine precision because all Hamiltonian elements are computed using exact algebraic expressions in MATLAB’s standard double-precision arithmetic. This ensures that the eigenvalues serve as a reliable benchmark for comparison with the -based results. In the approach, the spin projection is performed exactly, using Sanibel coefficients, as detailed above. This procedure avoids the numerical problems that could arise when using Löwdin’s projector or if the group-theoretical projection integral were discretized over a grid. However, adaptation inherently involves solving a GEP, which may introduce errors due to the conditioning of the overlap matrix and the limitations of the numerical solver. As noted in the Theory and Computational Details section, we verify that the rank of the overlap matrix corresponds to the full dimension of the symmetry subspace within a reasonable tolerance. Figure 3 demonstrates that the energy errors for all levels are negligibly small for practical purposes, confirming the reliability of our adaptation.
Figure 3.
Comparison of the energy differences from ED based on adaptation to or , as a function of the crystal momentum , for an , ring. The energy of the level is exact in both calculations ; therefore, is undefined. The numerical errors for degenerate irrep pairs, such as and , differ because the random selection of spin configurations for applying the projectors is performed independently for each sector.
To illustrate the utility of the spectrum, we follow Ref. [17] by focusing on two thermodynamic properties: magnetization and magnetic heat capacity, which are functions of the temperature and magnetic field. Both quantities can be readily computed using the complete set of eigenvalues. The thermal average of the magnetization is given by Equation (13):
where is the magnetic quantum number for state i, is the molecular partition function, , and represents the total energy, which includes the field-free energy obtained from ED and a Zeeman contribution that depends on the external magnetic field B, where g and denote the isotropic g-factor and the Bohr magneton, respectively.
The heat capacity , defined in Equation (14),
is expressed in terms of the internal energy and can be calculated from the thermal-energy fluctuation, as shown in Equation (15):
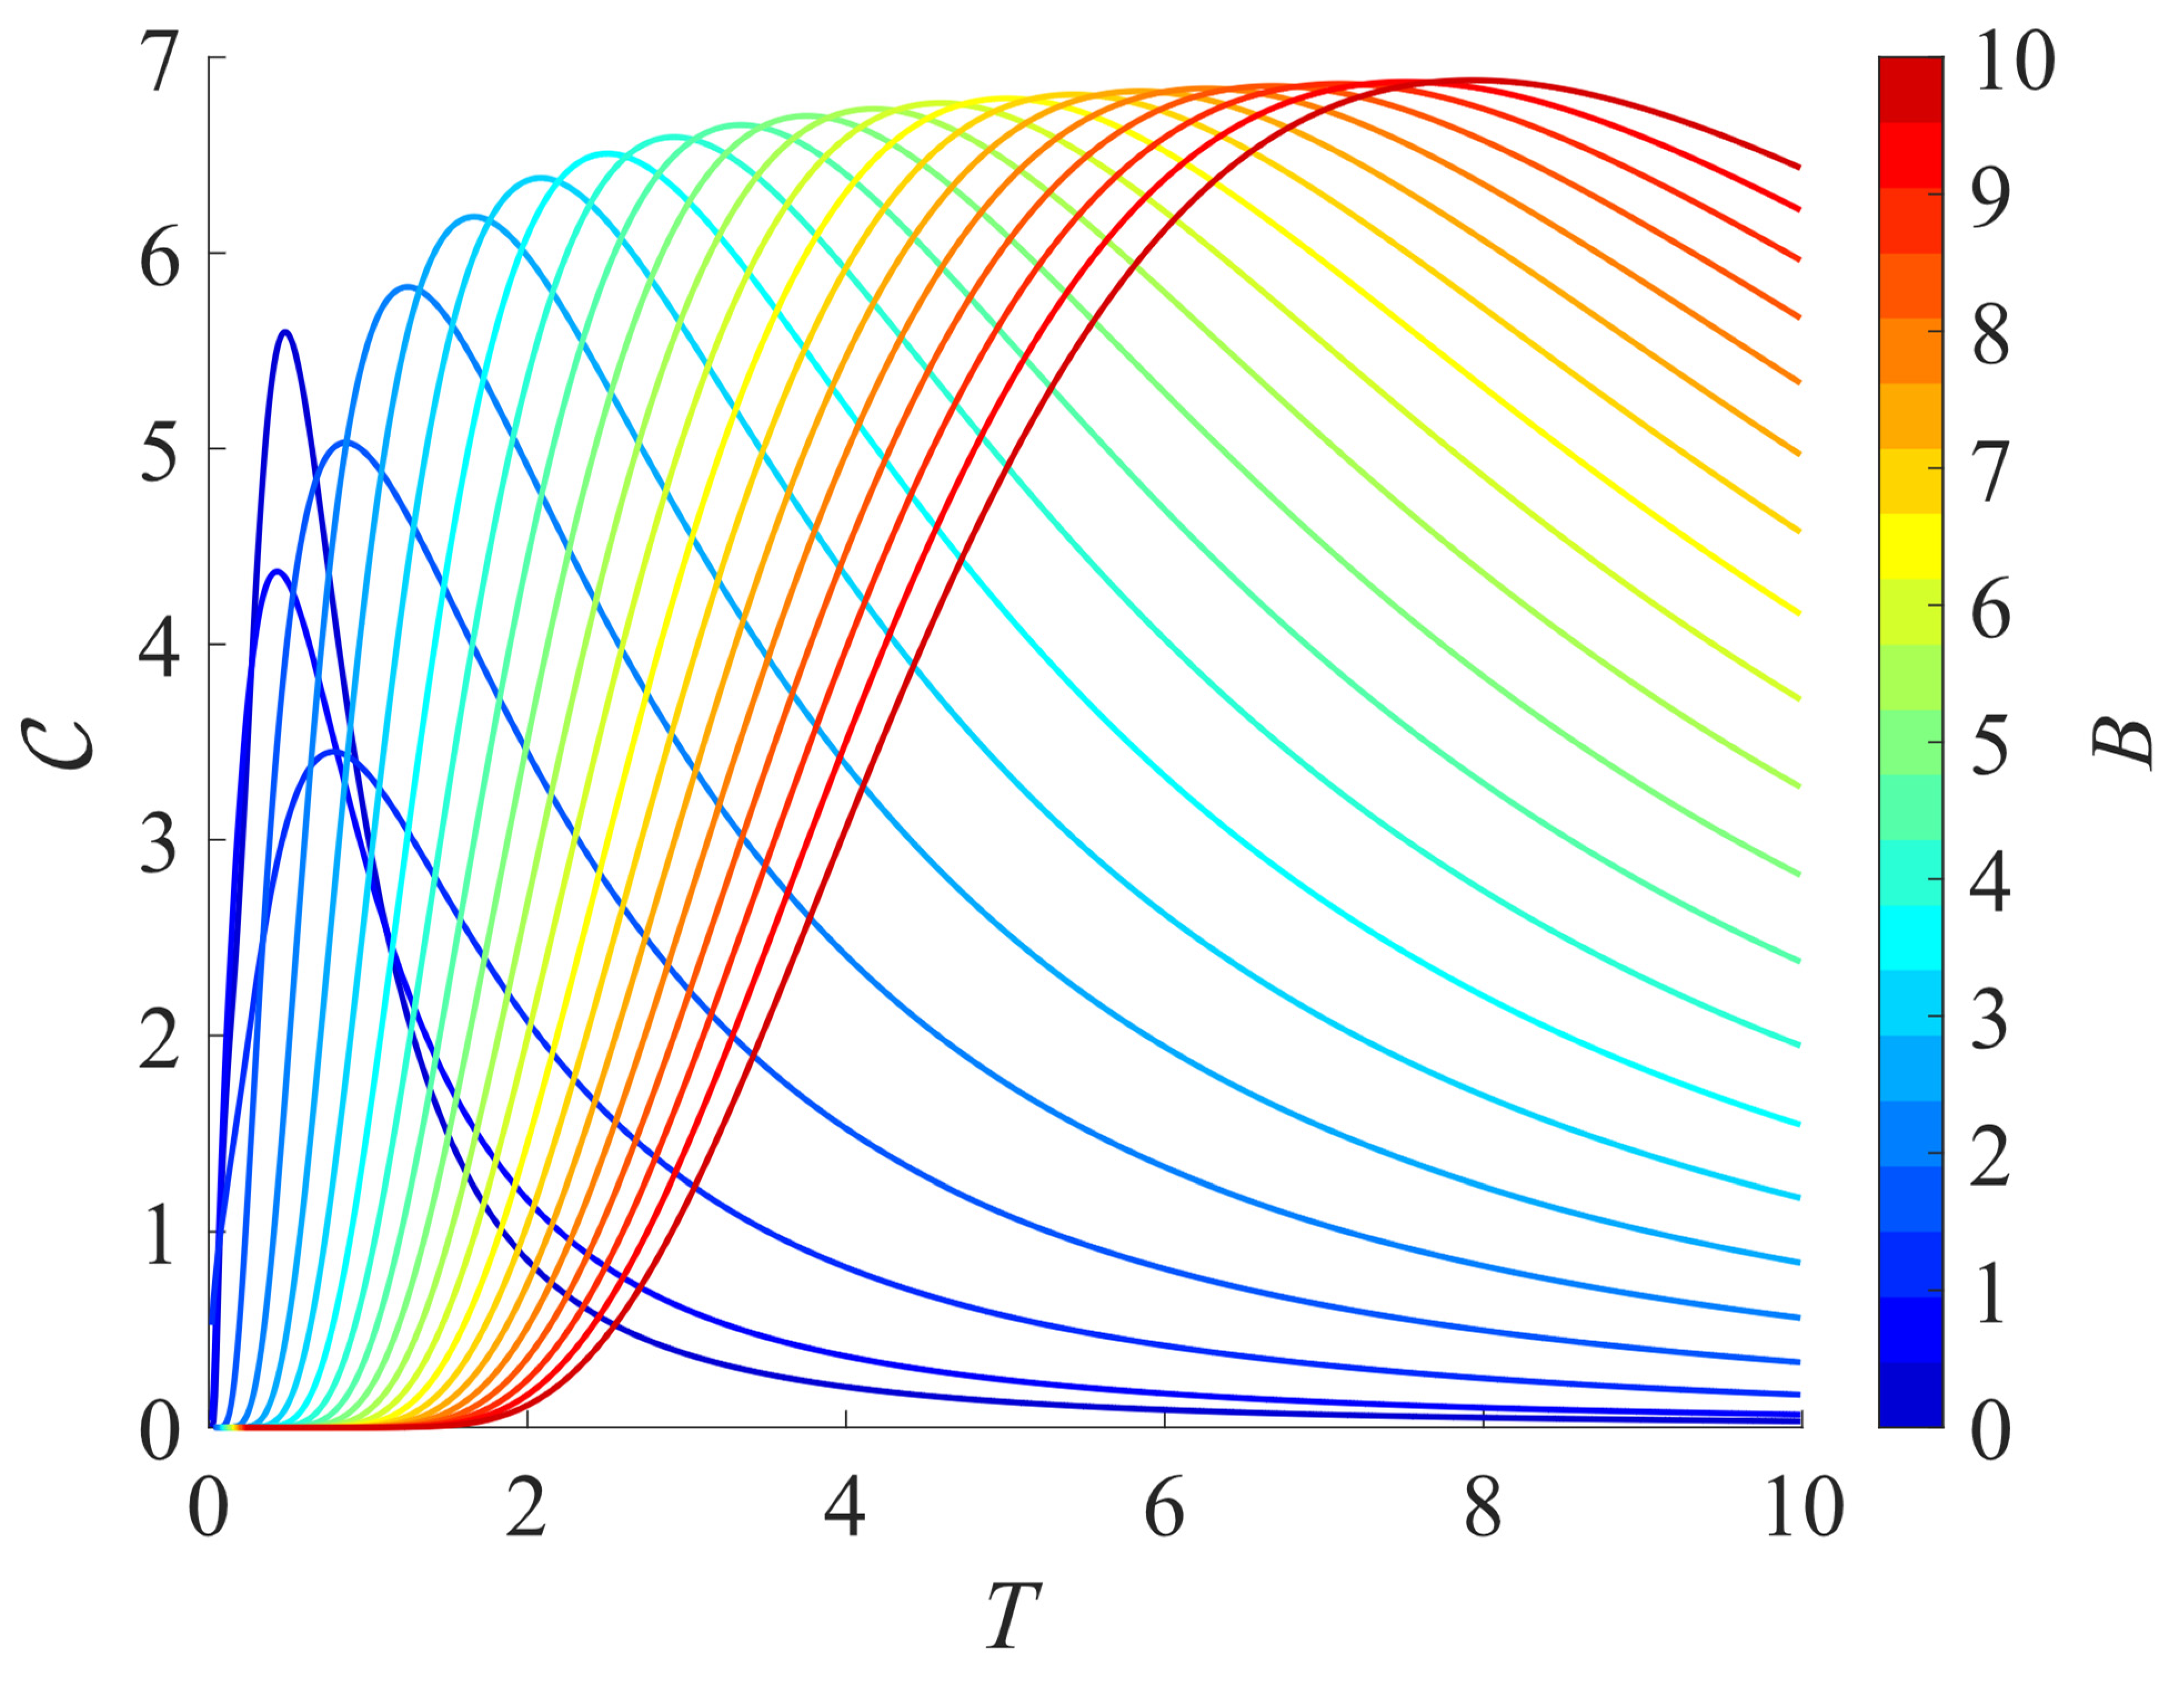
where . and are plotted in Figure 4 and Figure 5, respectively, for the , ring; we set and .
Figure 4.
Magnetization as a function of the external field B for various temperatures T (indicated by the color bar) for an , ring, with . Please refer to the main text for further details.
Figure 5.
Magnetic heat capacity of an antiferromagnetic , ring as a function of T for various field strengths B (indicated by the color bar).
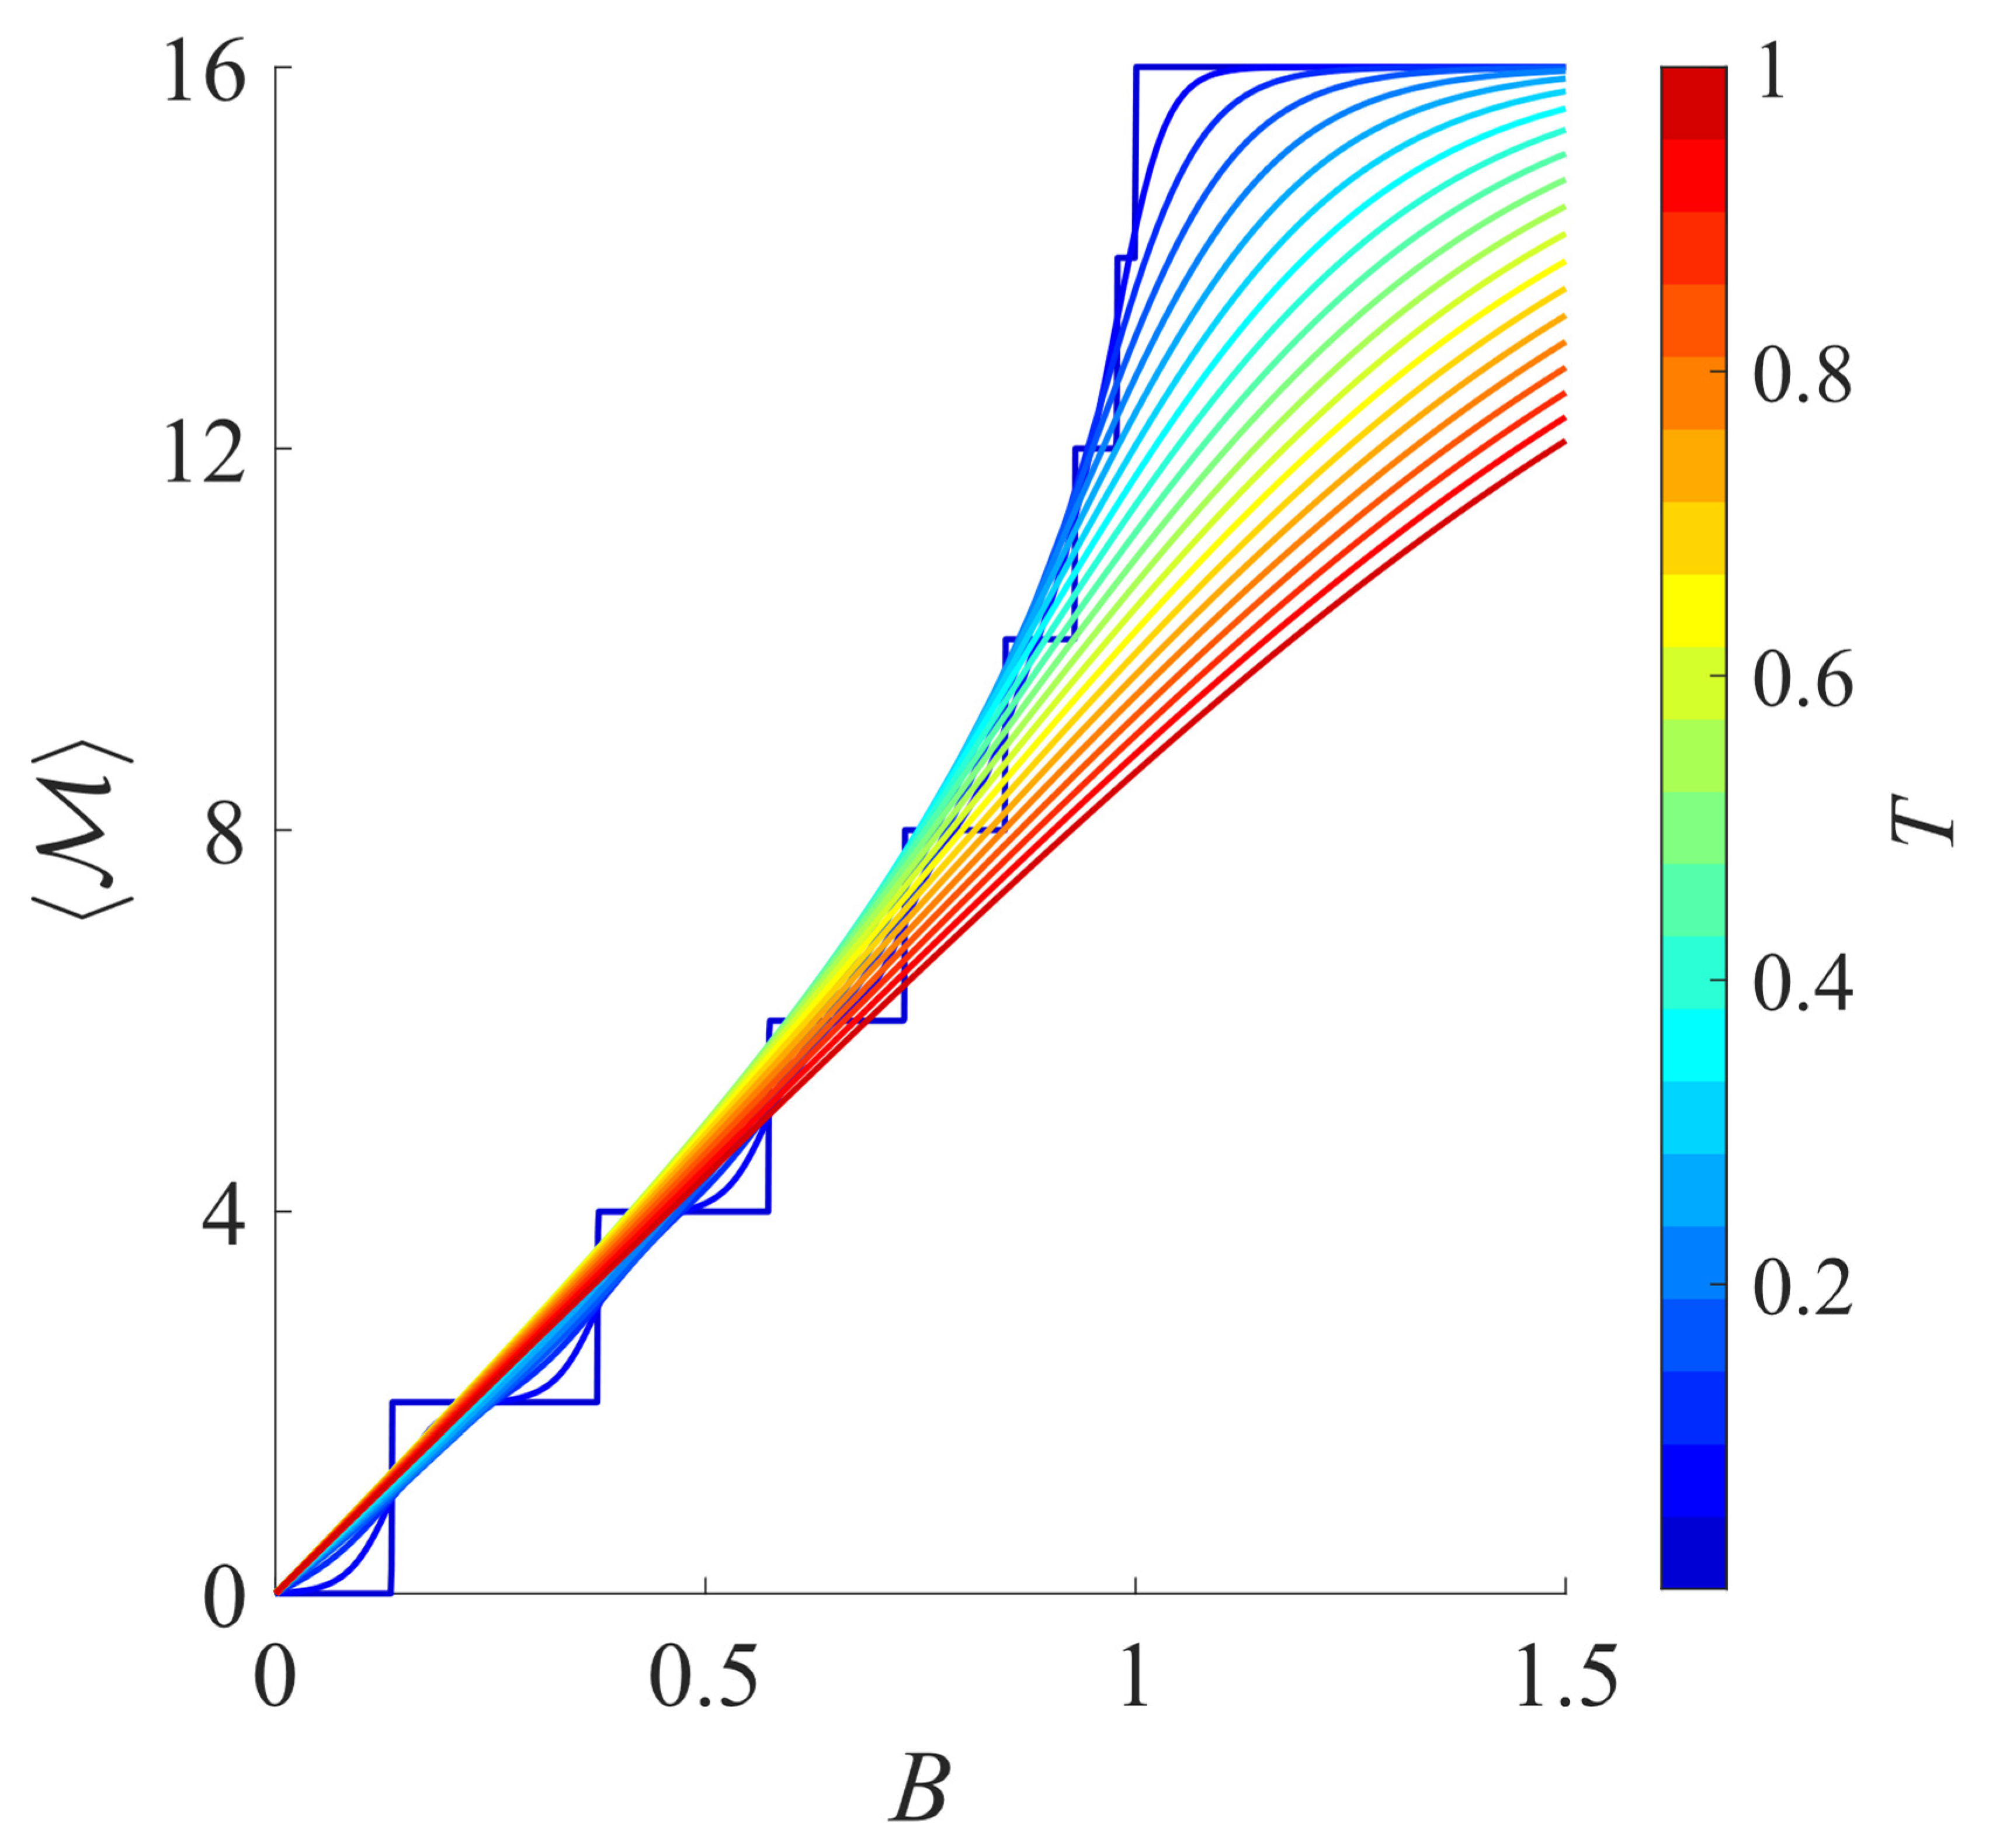
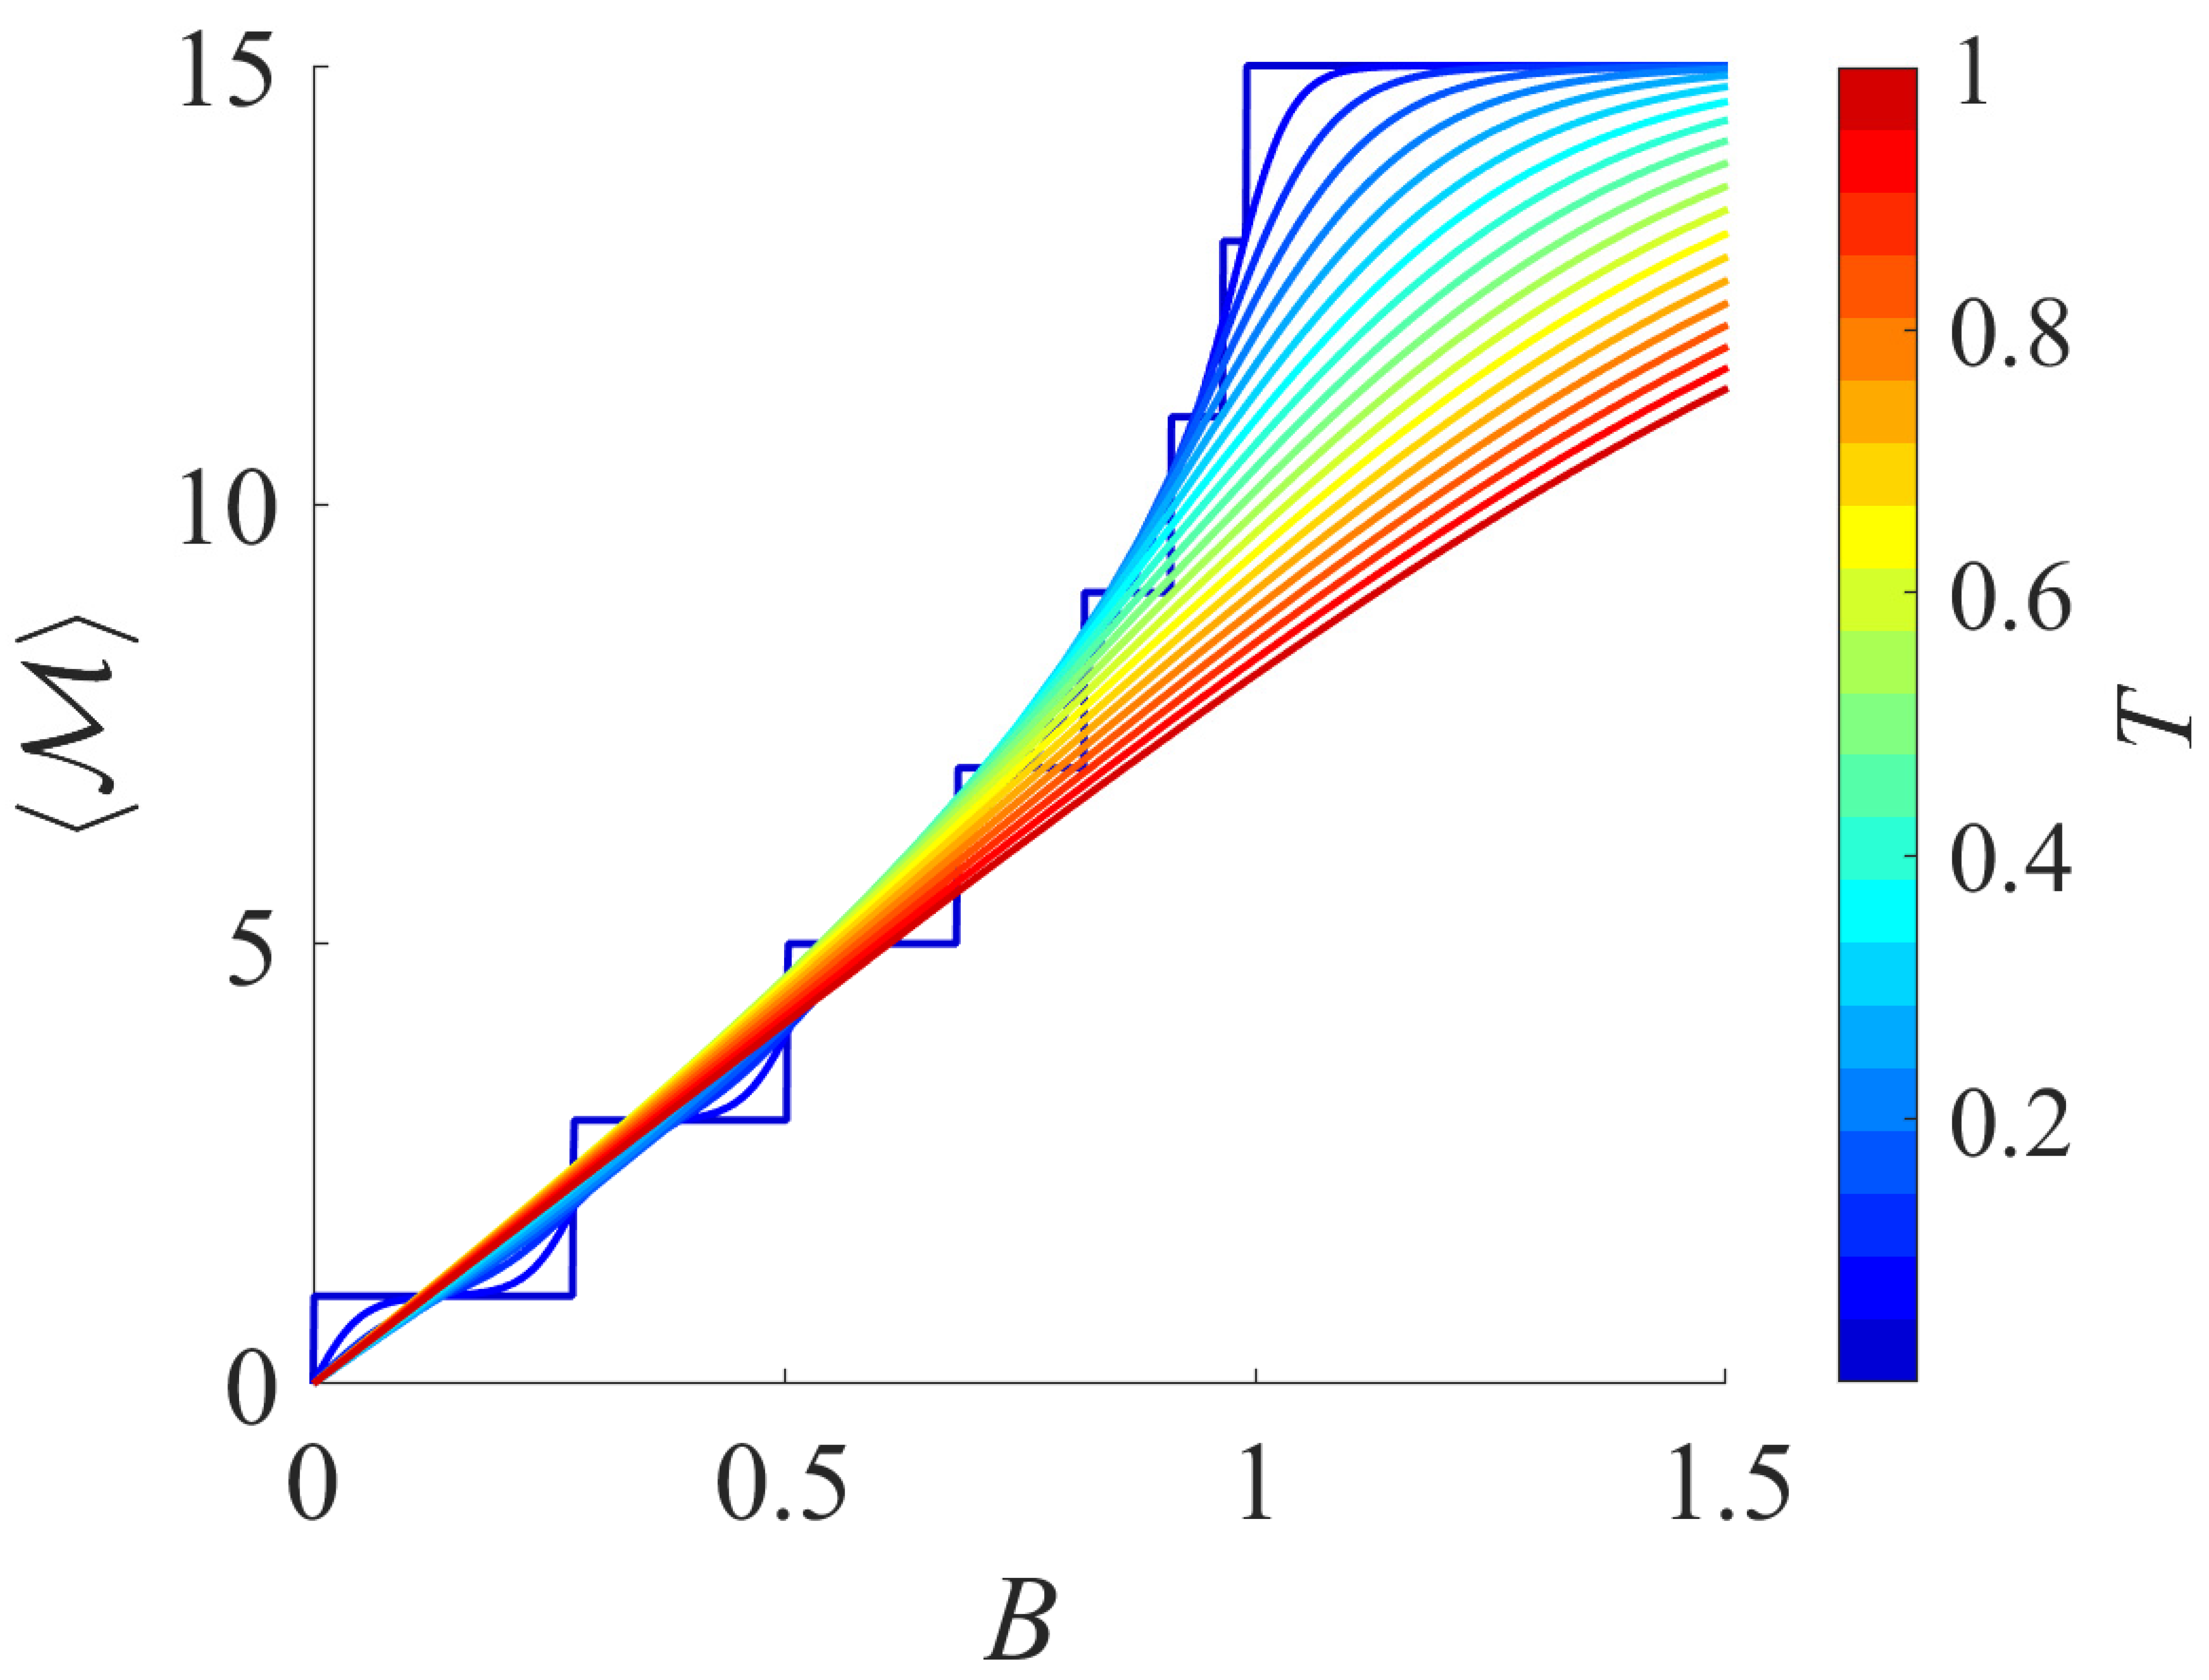
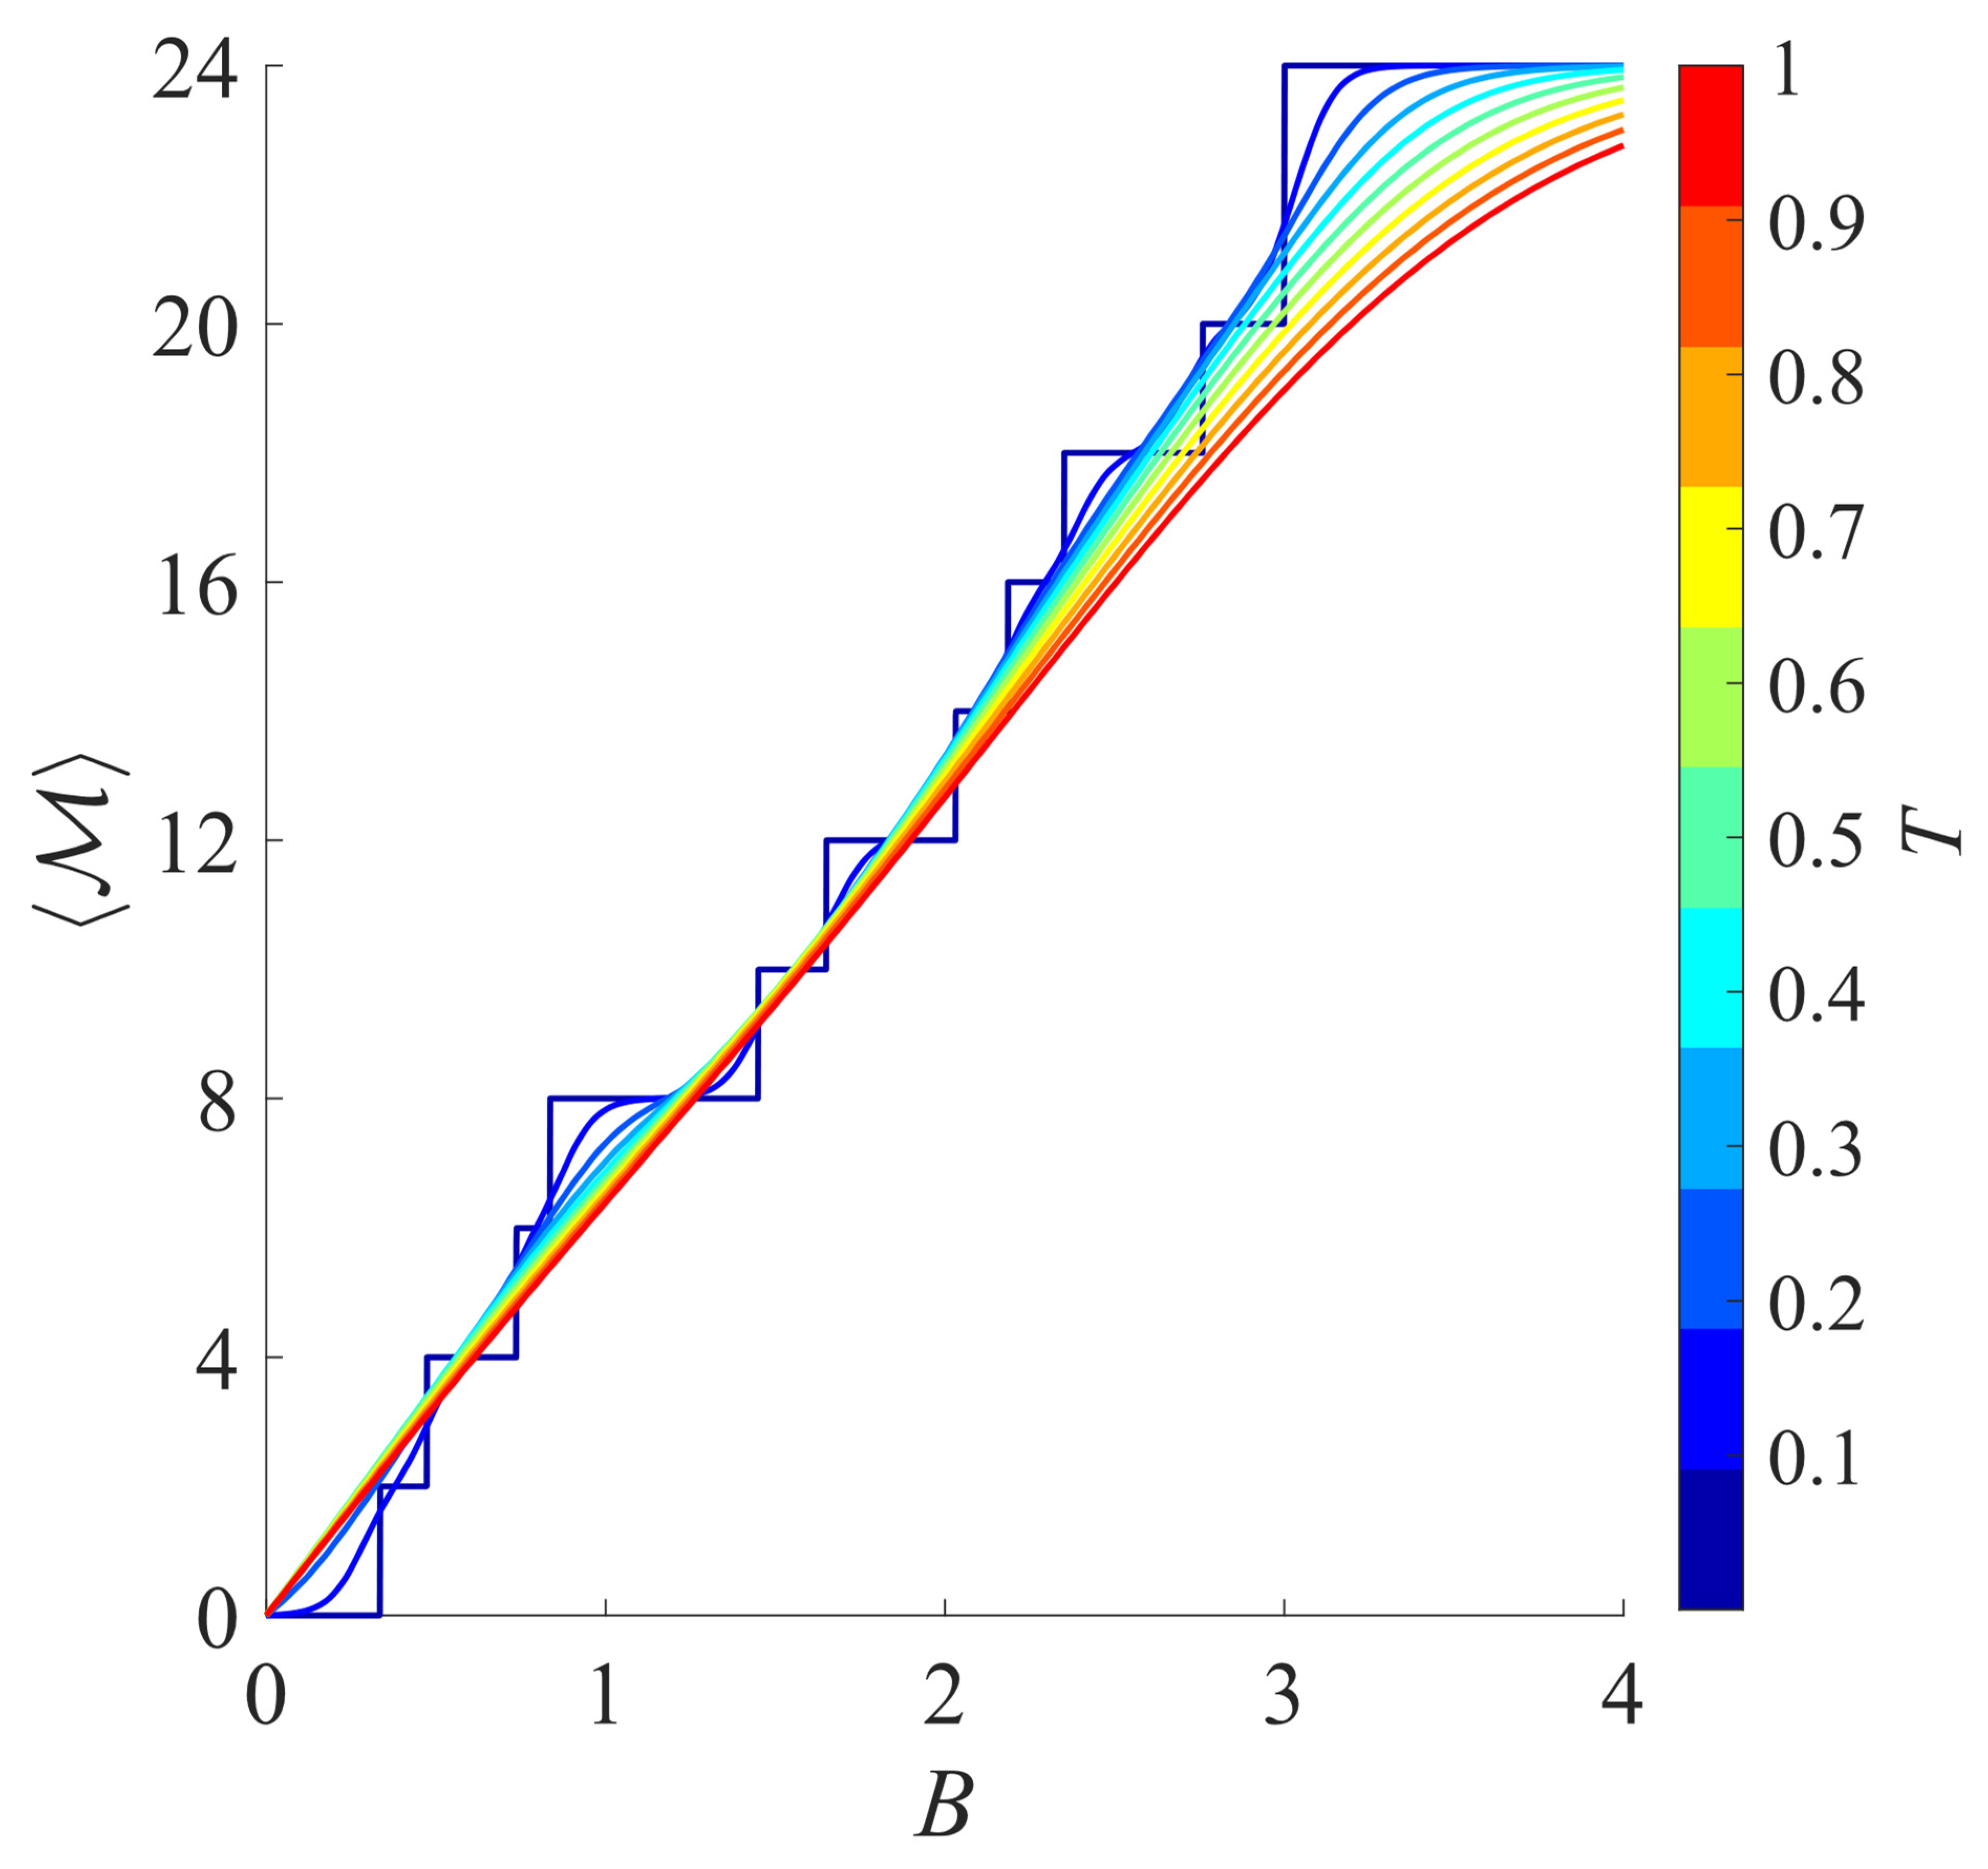
The magnetization of an , ring (a Kramers system, with half-integer spin) is non-zero at T = 0 in the zero-field limit; see Figure 6. In real systems at low temperatures, however, anisotropic effects—such as local zero-field splitting (ZFS) and anisotropic g-tensors—can also influence the magnetization.
Figure 6.
Magnetization as a function of B and T (color bar) for an , ring, with .
Icosahedron. By employing the point group and (and spin-flip) symmetries, low-energy spectra for various values of s have been obtained [33], and the icosahedron has been fully diagonalized by Konstantinidis [15]. However, full diagonalization for with such an approach would be challenging due to the significantly larger state-space dimension (cf. Table A1 in Appendix A). Although the Hamiltonian and overlap matrices in the basis are stored in a memory-efficient sparse format, exploiting this advantage for complete diagonalization would require specialized libraries. The built-in eig function in MATLAB, which computes a full spectrum for (generalized) eigenvalue problems, does not accept sparse inputs; sparse matrices must thus be converted into their full counterparts. On the other hand, the iterative solver eigs is designed to compute only a subset of the eigenvalues (and eigenvectors) of sparse matrices.
Using the GC method, it took three days on 128 processors to construct the Hamiltonian in the largest subspace in the icosahedron (dimension 3315, , ), primarily due to the computationally demanding transformations between coupling schemes [10]. Using our current projection method on a desktop computer with six CPU cores, setting up the GEP in the same space (as explained in the Theory and Computational Details section, we separate the five components of , reducing the matrix dimensions in the , space to ; see Table A2 in Appendix A) took only a few seconds, and determining the full spectrum took two minutes (this estimate is provided primarily for illustrative purposes, as it is likely possible to improve the implementation of our method to reduce the runtime). In our current implementation, the main bottleneck encountered is the construction of spin eigenstates, which requires evaluating the analytical form of the group-theoretical integral of Equation (7). The spin adaptation for is more demanding for systems of similar Hilbert-space dimensions because the integral—as explained in Ref. [26]—decomposes into a sum of analytically evaluable standard integrals, and the number of terms in the sum grows steeply as s increases. In contrast, for systems, only a single standard integral needs to be evaluated.
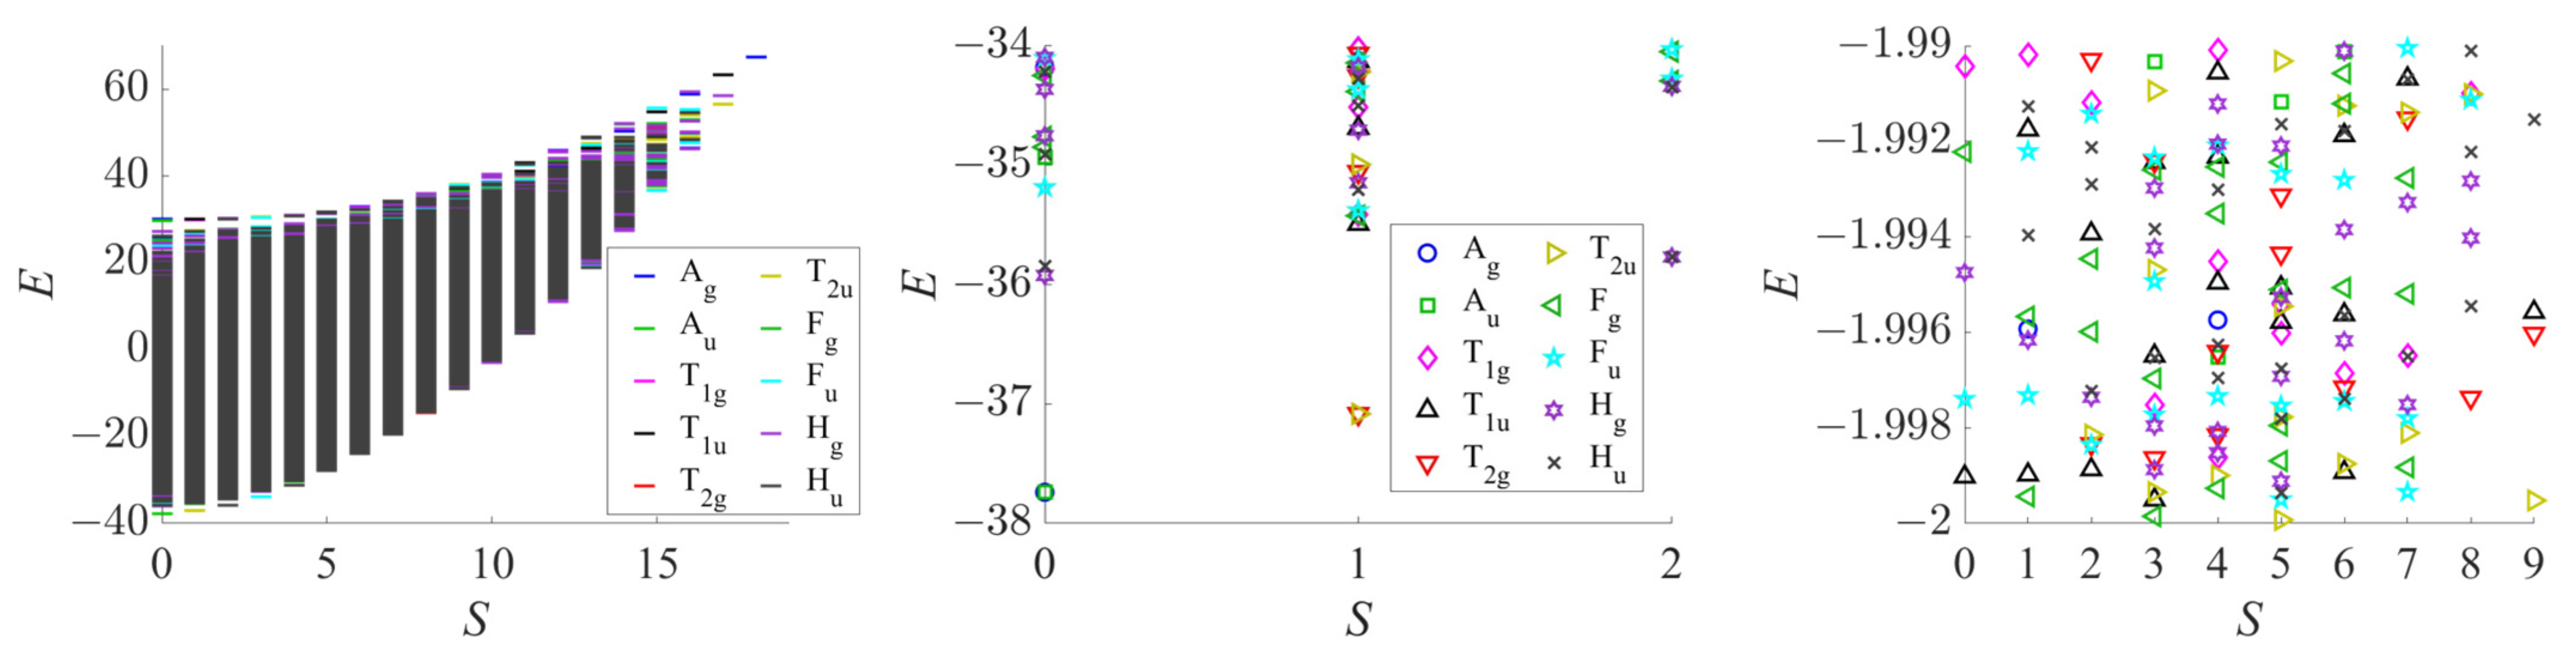
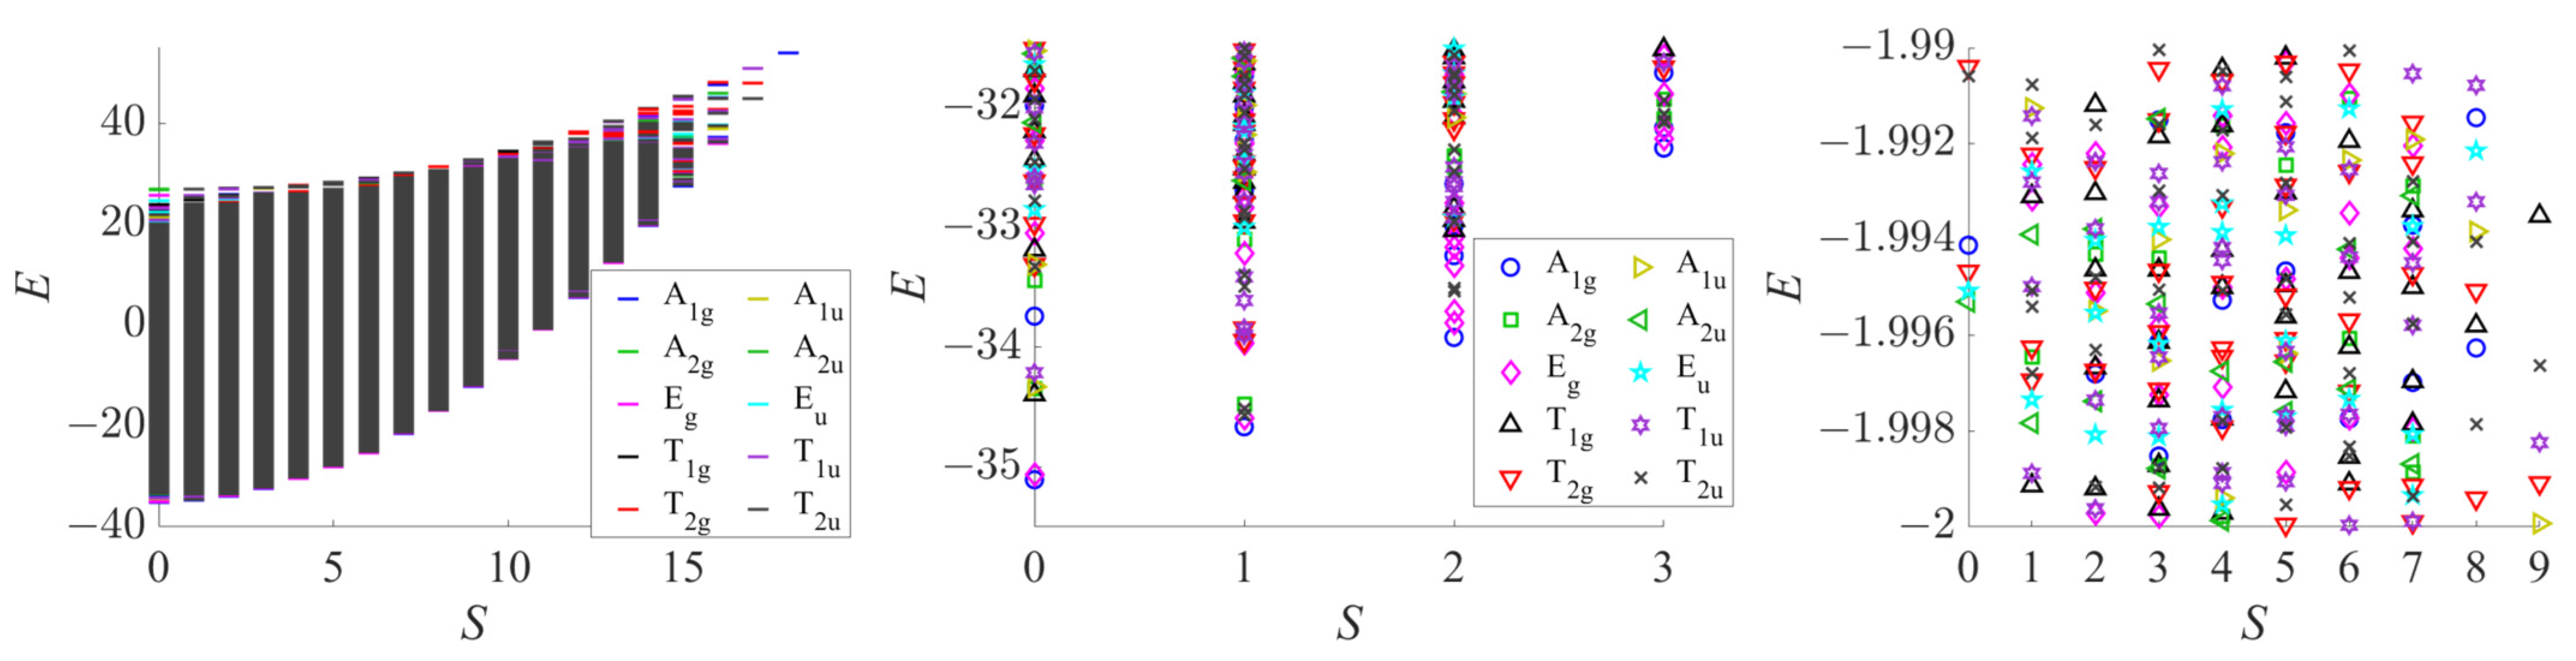
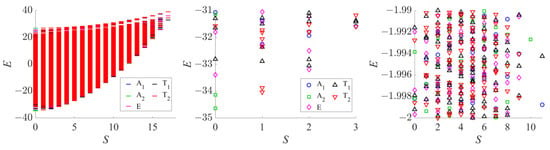
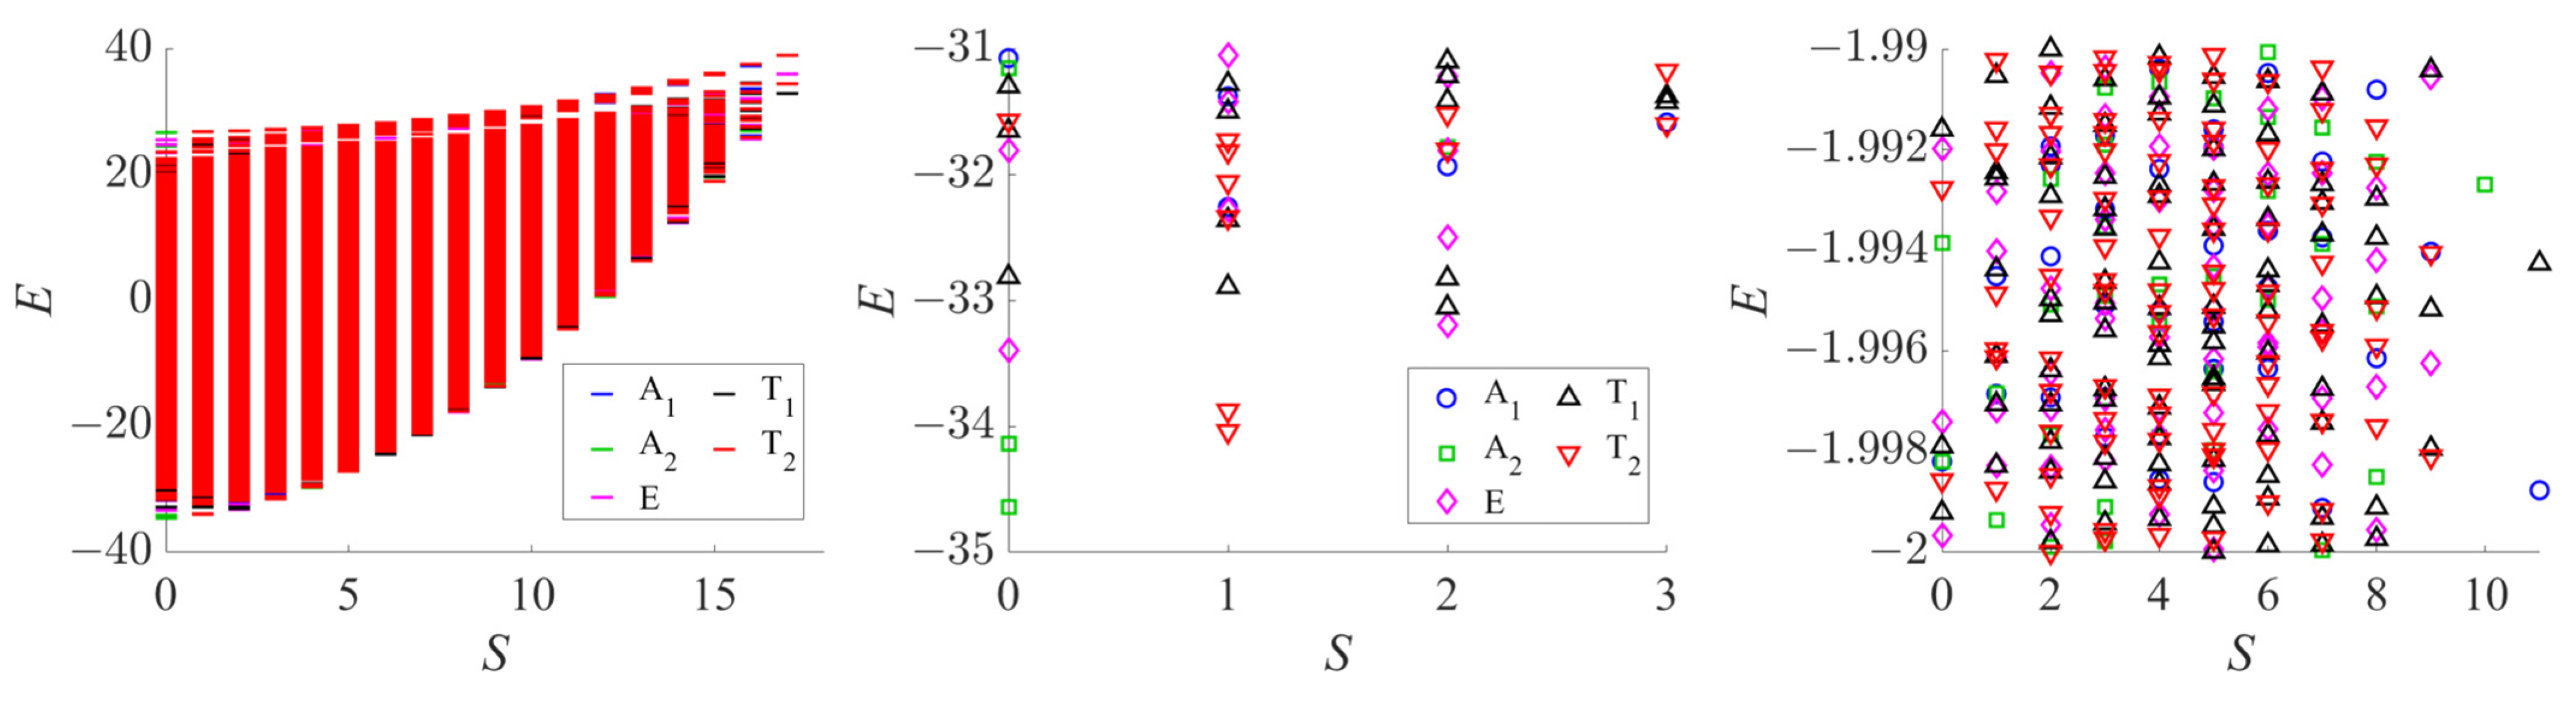
Even with 256 processors, a full adaptation with respect to the total spin and symmetry for the icosahedron could not be completed using the GC approach [10]. Instead, this system was diagonalized by adopting the subgroup of , which permits the construction of a compatible coupling scheme; however, the Hamiltonian blocks are significantly larger compared to full symmetry. With our approach, we are now able to adapt to within the complete point group without any difficulty: the determination of the full spectrum of the icosahedron took us approximately 16 h on a cluster node with 64 GB RAM and eight CPU cores. The dimensions of the respective subspaces are collected in Table A3 and spectra are illustrated in Figure 7. If only spin symmetry were used (as is done, e.g., in the MAGPACK program [34]) instead of combined spin and PG symmetry, then the matrices are considerably larger and the relevant dimensionrs can be derived from the table in the Appendix A. For example, for S = 4, if one takes into account the degrees of degeneracy of the PG irreps, this results in a prohibitive matrix dimension of 258,192, according to Equation (16).
Figure 7.
The full spectrum of the icosahedron (left), the low-energy section (middle), and an arbitrary section from its intermediate energy range (right).
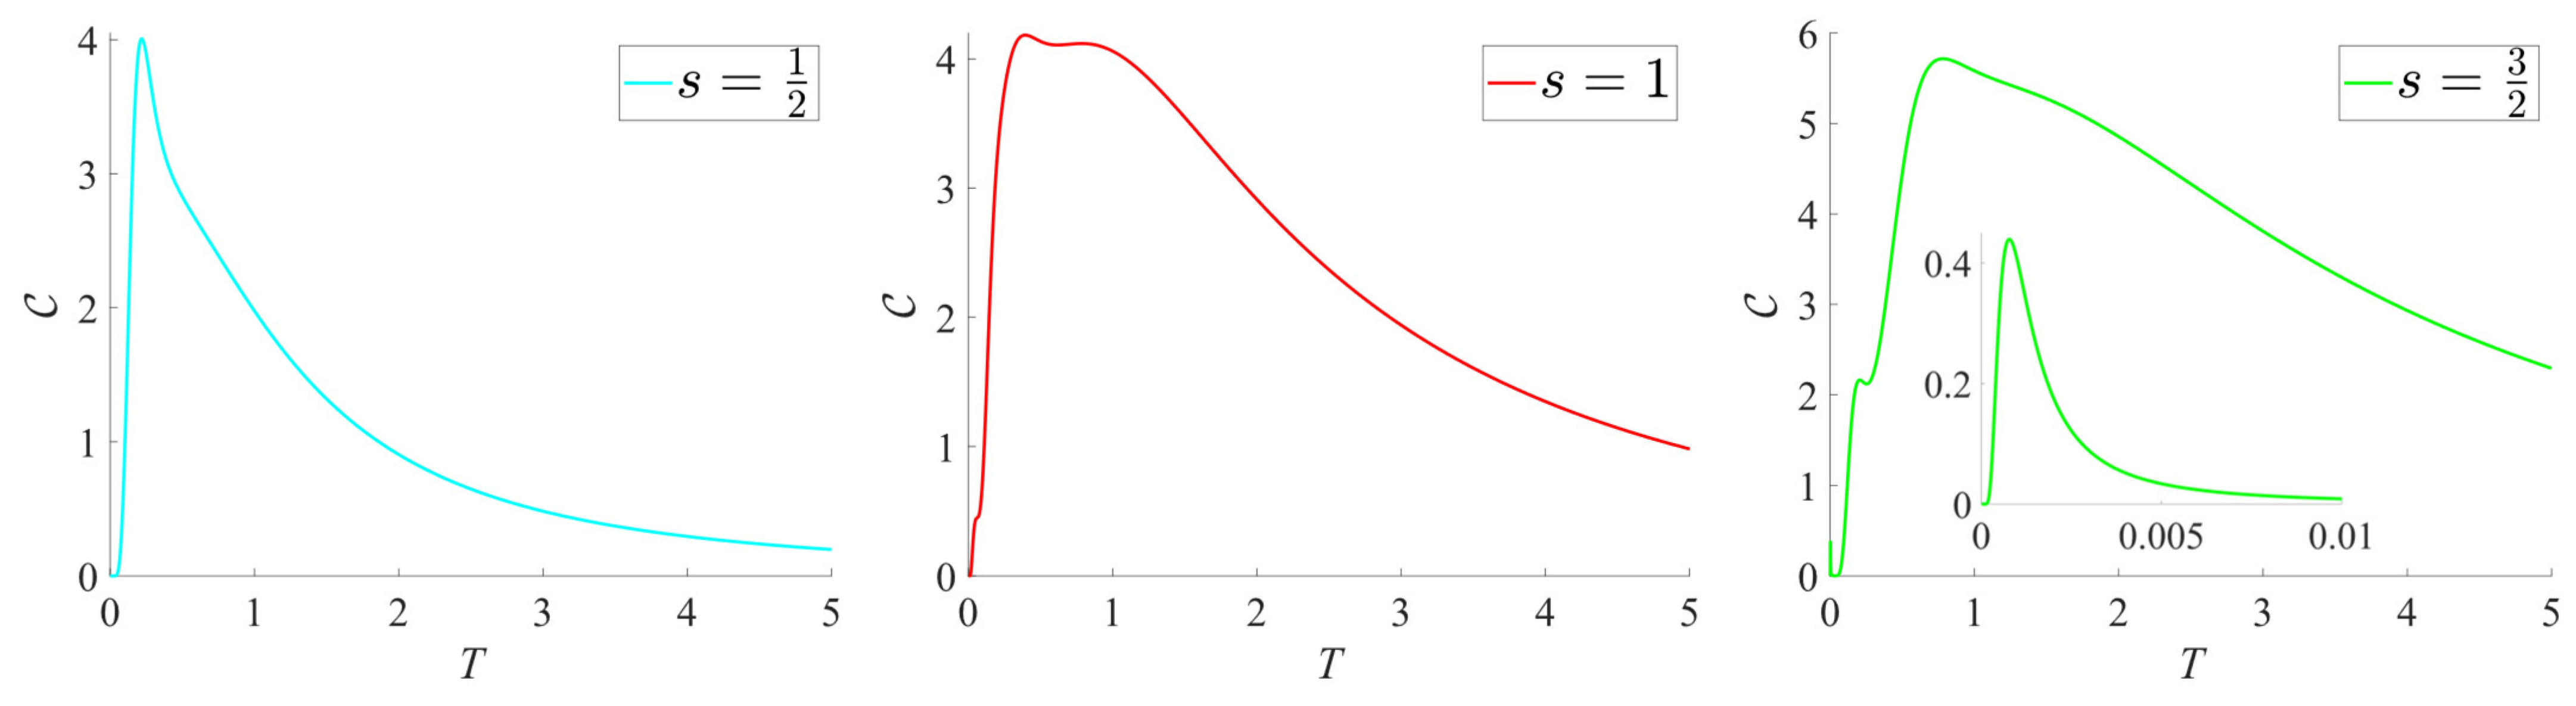
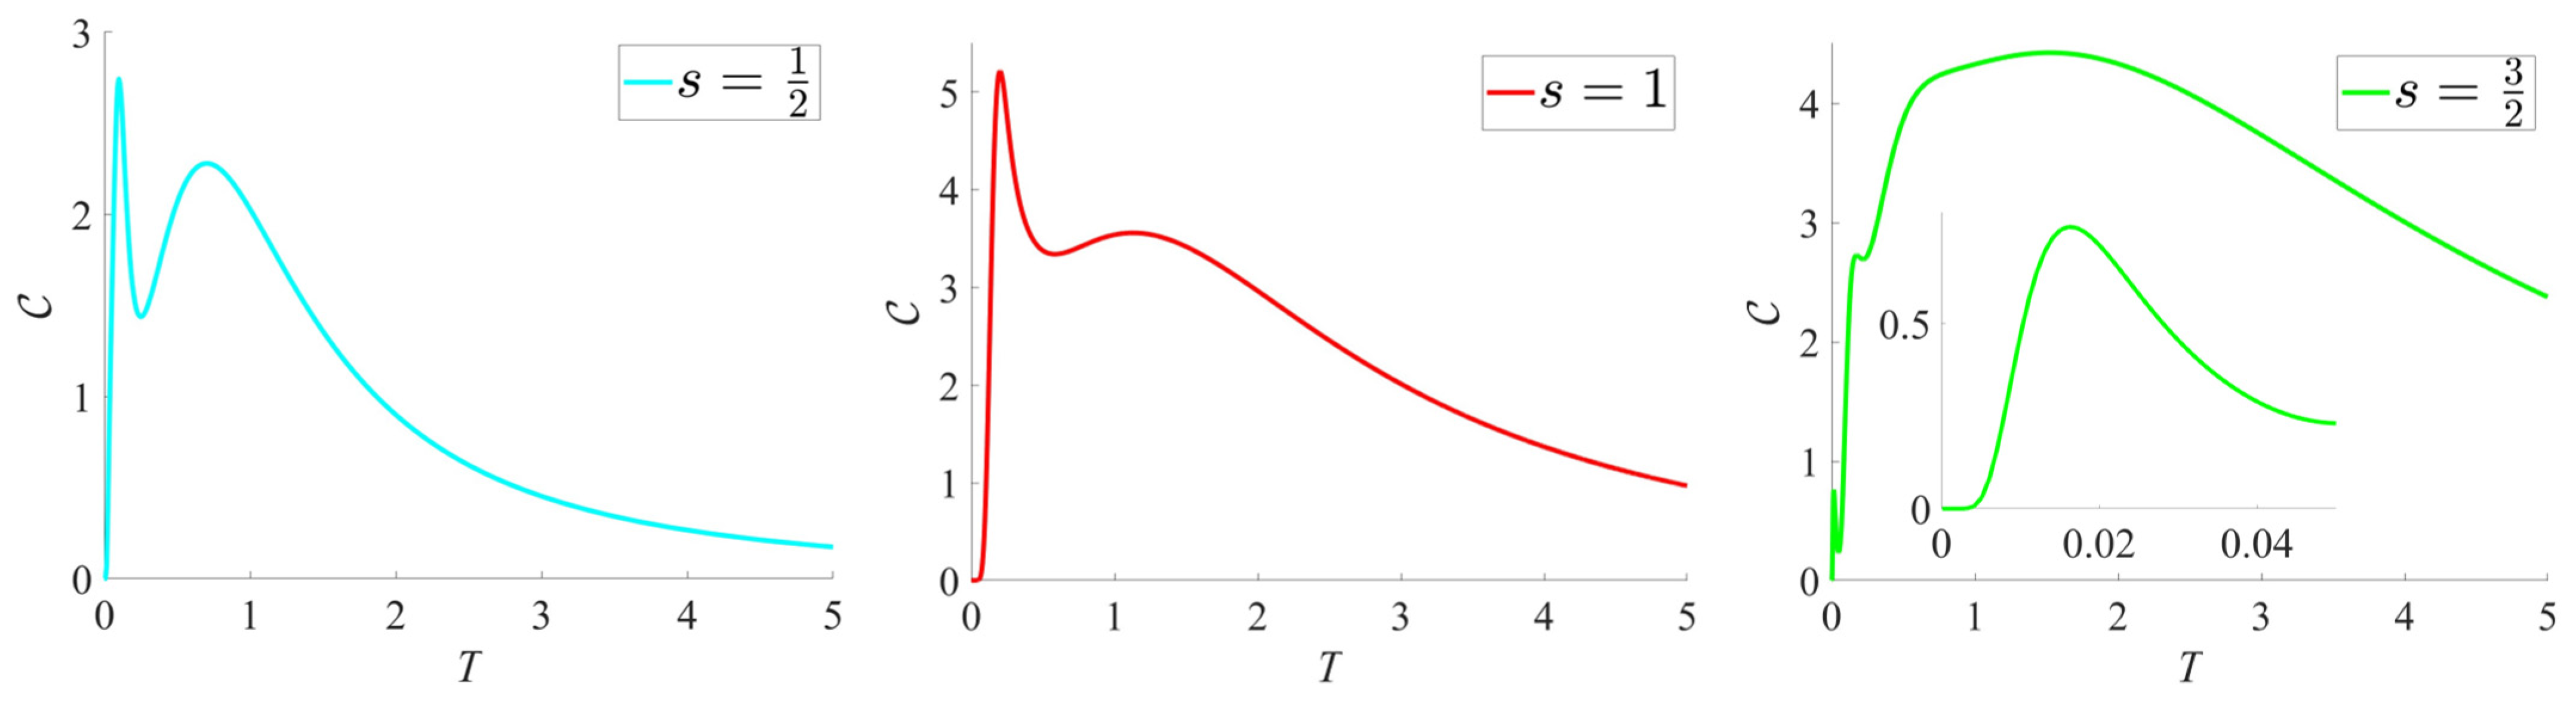
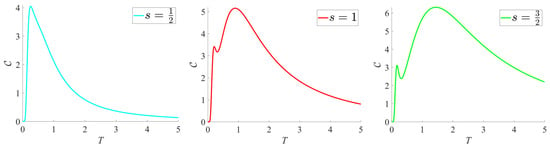
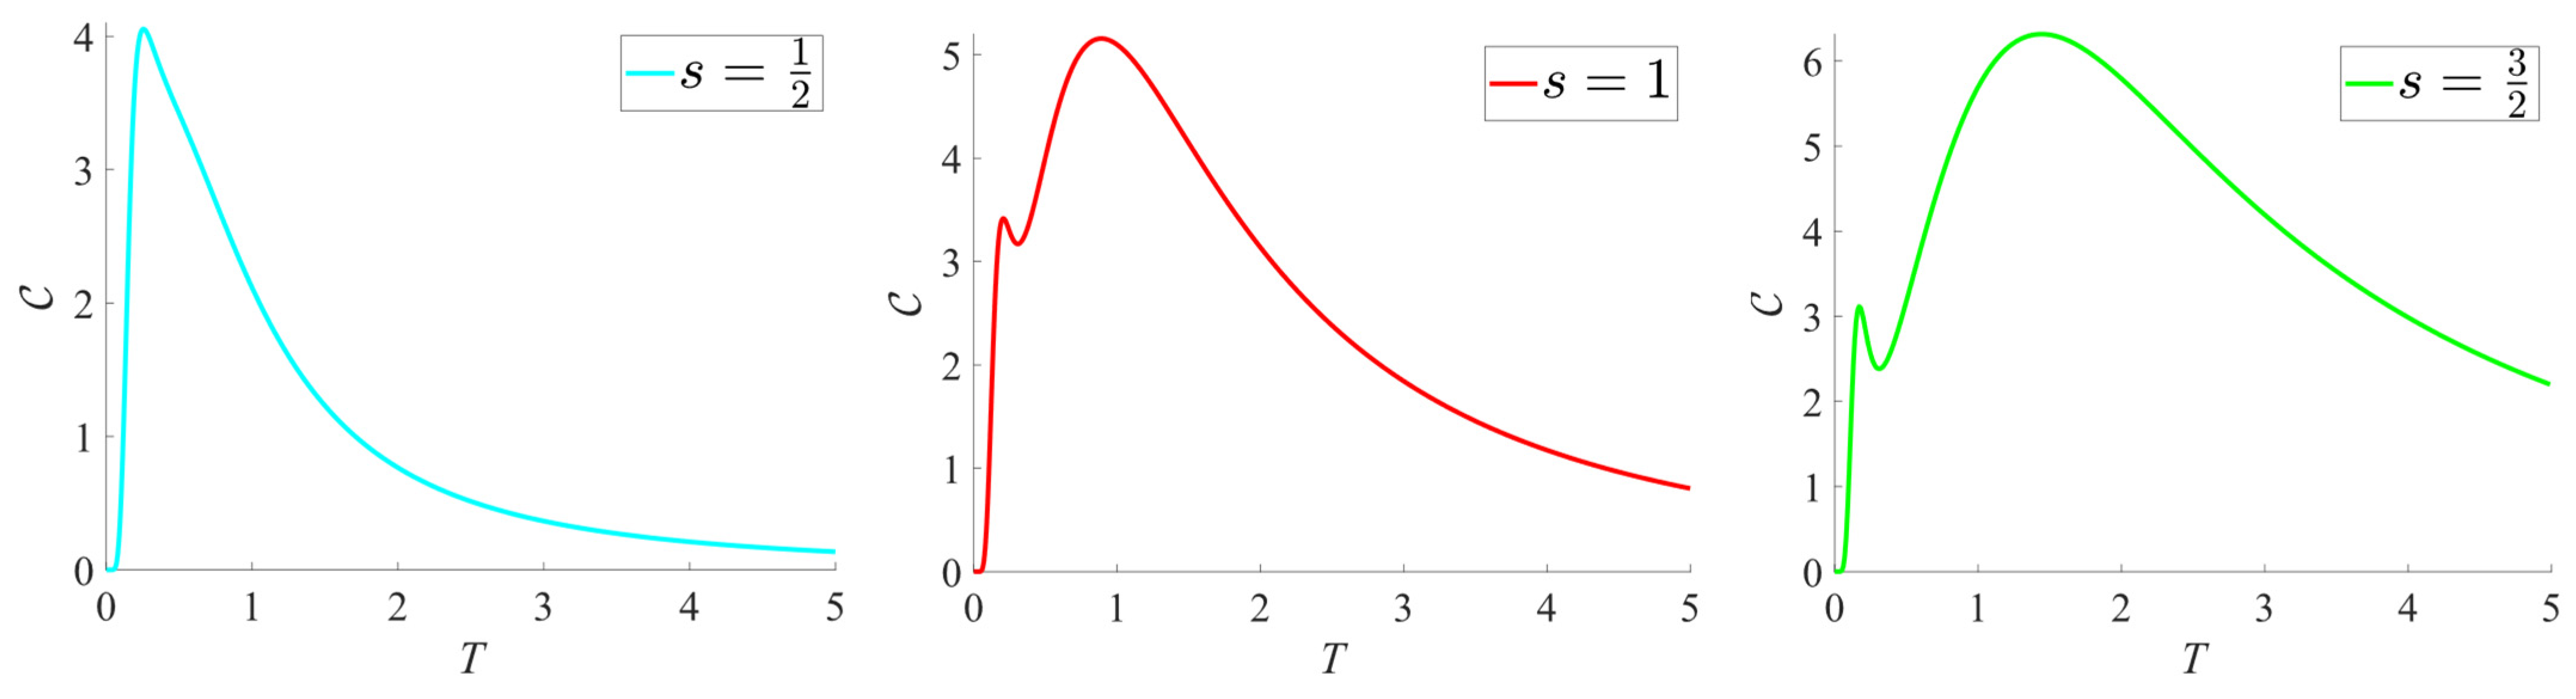
The magnetic heat capacities in the zero field for icosahedrons with , , and are shown in Figure 8. Note that for and heat capacity values have also been plotted in Ref. [15] (albeit with a logarithmic temperature axis). There are two peaks in the low-temperature part of the curve for the system. The first peak at is due to a near-degeneracy of the ground state, as the lowest levels in sectors and have energies of and , respectively. We verified the curve for s = 1/2 with exact diagonalization and without using symmetries, and the curves for s = 1 and s = 3/2 were confirmed through FTLM calculations employing only symmetry [35].
Figure 8.
Zero-field magnetic heat capacity for an icosahedron with , , and . A low-temperature peak is observed at for (inset), reflecting the near-degeneracy of this system’s ground state.
Cuboctahedron. A comparison between diagonalization in the and spaces for the cuboctahedron, as shown in Figure 9, again demonstrates the good agreement of our approach. The magnetization is plotted as a function of B and T in Figure 10.
Figure 9.
Comparison of energy differences in diagonalization based on adaptation to or , as a function of the irreps of the point group, for an cuboctahedron.
Figure 10.
Magnetization as a function of B and T for an cuboctahedron.
The system was solved by Schnalle and Schnack in the compatible subgroup, while only the low-energy part of the spectrum was fully resolved with respect to [10,19]. Here, we have for the first time obtained a full symmetry classification of the entire spectrum (the subspace dimensions are given in Table A4 in Appendix A); see Figure 11.
Figure 11.
The full spectrum of the cuboctahedron (left), the low-energy section (middle), and an arbitrary section from its intermediate energy range (right).
We plot the magnetic heat capacities of the cuboctahedron in Figure 12. A comparison with Ref. [10] reveals that our calculated curves coincide with theirs if the temperature axis is scaled by a factor of two. This difference could arise from a distinct choice of conventions in defining the exchange constant J in the Heisenberg Hamiltonian. However, our convention for energies matches those in Ref. [10], as is apparent from Figure 5 and Figure 6 in that work, implying an inconsistency in the treatment of J-conventions [35].
Figure 12.
Zero-field magnetic heat capacity for a cuboctahedron with , , and . A low-temperature peak at for (inset) is due to the near-degeneracy of the ground state of this cuboctahedron (cf. Figure 10).
Truncated Tetrahedron. Finally, we consider the truncated tetrahedron, which, to the best of our knowledge, has never been fully diagonalized for . This may be attributed to the fact that the relevant point group (which is isomorphic to the octahedral group ), with only five irreps comprising a total of 10 components, is smaller than or , which have 10 irreps each and 20 and 30 components, respectively. Consequently, the matrices involved are significantly larger but still manageable if the full symmetry is exploited (see Table A5 in Appendix A). The spectrum of the truncated tetrahedron is presented in Figure 13, and the heat capacities of the , , and systems are shown in Figure 14.
Figure 13.
The full spectrum of the truncated tetrahedron (left), the low-energy section (middle), and an arbitrary section from the intermediate energy range (right).
Figure 14.
Zero-field magnetic heat capacity for the truncated tetrahedron for , , and .
4. Summary and Conclusions
We have presented a simple and efficient approach for the exact diagonalization (ED) of isotropic spin clusters by combining spin and point-group (PG) symmetries. This method applies projection operators to uncoupled basis states, thereby eliminating the need for the complex recoupling transformations typically required in schemes based on genealogical coupling (GC). The accuracy and versatility of the present projection approach, and its ability to handle systems that are computationally inaccessible with other techniques, was demonstrated on spin rings and polyhedra. Notably, we could fully determine and classify the spectra of the icosahedron and cuboctahedron, which had previously been only partially resolved due to the computational limitations imposed by recoupling transformations, which are avoided in the present projection approach. Moreover, we diagonalized the truncated tetrahedron for the first time, a task that would pose a significant challenge without the full exploitation of symmetry. Due to its numerical efficiency, conceptual simplicity, and straightforward implementation, this projection method potentially represents a standard for tackling large Heisenberg spin clusters via exact diagonalization.
Author Contributions
Conceptualization, S.G.T.; methodology, S.G.T.; investigation, S.G.T.; writing—original draft preparation, S.G.T.; writing—review and editing, S.G.T. and T.D.K.; supervision, T.D.K. All authors have read and agreed to the published version of the manuscript.
Funding
This work was supported by the Deutsche Forschungsgemeinschaft (DFG) under Project 535298924.
Data Availability Statement
Data are contained within the article.
Acknowledgments
We thank J. Schnack for providing FTLM results and TU Berlin for their computational resources.
Conflicts of Interest
The authors declare no conflicts of interest.
Appendix A
Here, we compile the dimensions of symmetry subspaces for some of the systems discussed in this work, specifically the icosahedron, cuboctahedron, and truncated tetrahedron.
Table A1.
Dimensions of the combined and point-group subspaces for the icosahedron.
Table A2.
Dimensions of the combined total spin and point-group subspaces for the s = 1 icosahedron.
Table A3.
Dimensions of the combined total spin and point-group subspaces for the icosahedron.
Table A4.
Dimensions of the combined total spin and point-group subspaces for the cuboctahedron.
Table A5.
Dimensions of the combined total spin and point-group subspaces for the truncated tetrahedron.
References
- Gatteschi, D.; Sessoli, R.; Villain, J. Molecular Nanomagnets; Oxford University Press: Oxford, UK, 2006. [Google Scholar]
- Chilton, N.F. Molecular Magnetism. Annu. Rev. Mater. Res. 2022, 52, 79–101. [Google Scholar] [CrossRef] [Scilit]
- Bencini, A.; Gatteschi, D. Electron Paramagnetic Resonance of Exchange Coupled Systems; Springer: Berlin/Heidelberg, Germany, 1990. [Google Scholar]
- Schnack, J.; Ummethum, J. Advanced quantum methods for the largest magnetic molecules. Polyhedron 2013, 66, 28–33. [Google Scholar] [CrossRef] [Scilit]
- Levitt, M.H. Spin Dynamics; John Wiley & Sons: Hoboken, NJ, USA, 2008. [Google Scholar]
- Waldmann, O. Symmetry and energy spectrum of high-nuclearity spin clusters. Phys. Rev. B 2000, 61, 6138. [Google Scholar] [CrossRef] [Scilit]
- Tsukerblat, B.S. Group Theory in Chemistry and Spectroscopy; Courier Corporation: North Chelmsford, MA, USA, 2006. [Google Scholar]
- Klemm, R.A.; Efremov, D.V. Single-ion and exchange anisotropy effects and multiferroic behavior in high-symmetry tetramer single-molecule magnets. Phys. Rev. B 2008, 77, 184410. [Google Scholar] [CrossRef] [Scilit]
- Ghassemi Tabrizi, S.; Arbuznikov, A.V.; Kaupp, M. Exact Mapping from Many-Spin Hamiltonians to Giant-Spin Hamiltonians. Chem. Eur. J. 2018, 24, 4689–4702. [Google Scholar] [CrossRef] [Scilit]
- Schnalle, R.; Schnack, J. Calculating the energy spectra of magnetic molecules: Application of real- and spin-space symmetries. Int. Rev. Phys. Chem. 2010, 29, 403–452. [Google Scholar] [CrossRef] [Scilit]
- Ghassemi Tabrizi, S. Point-group selection rules and universal momentum-transfer dependencies for inelastic neutron scattering on molecular spin clusters. Phys. Rev. B 2021, 103, 214422. [Google Scholar] [CrossRef] [Scilit]
- Ghassemi Tabrizi, S. Triaxial magnetic anisotropy in the two-dimensional ferromagnetic semiconductor CrSBr. Phys. Rev. B 2021, 104, 144416. [Google Scholar]
- Waldmann, O. Q dependence of the inelastic neutron scattering cross section for molecular spin clusters with high molecular symmetry. Phys. Rev. B 2003, 68, 174406. [Google Scholar] [CrossRef] [Scilit]
- Raghu, C.; Rudra, I.; Sen, D.; Ramasesha, S. Properties of low-lying states in some high-nuclearity Mn, Fe, and V clusters: Exact studies of Heisenberg models. Phys. Rev. B 2001, 64, 064419. [Google Scholar] [CrossRef] [Scilit]
- Konstantinidis, N.P. Antiferromagnetic Heisenberg model on clusters with icosahedral symmetry. Phys. Rev. B 2005, 72, 064453. [Google Scholar] [CrossRef] [Scilit]
- Schnack, J.; Schulenburg, J.; Richter, J. Magnetism of the N = 42 kagome lattice antiferromagnet. Phys. Rev. B 2018, 98, 094423. [Google Scholar] [CrossRef] [Scilit]
- Heitmann, T.; Schnack, J. Combined use of translational and spin-rotational invariance for spin systems. Phys. Rev. B 2019, 99, 134405. [Google Scholar]
- Delfs, C.; Gatteschi, D.; Pardi, L.; Sessoli, R.; Wieghardt, K.; Hanke, D. Magnetic properties of an octanuclear iron(III) cation. Inorg. Chem. 1993, 32, 3099–3103. [Google Scholar] [CrossRef] [Scilit]
- Schnalle, R.; Schnack, J. Numerically exact and approximate determination of energy eigenvalues for antiferromagnetic molecules using irreducible tensor operators and general point-group symmetries. Phys. Rev. B 2009, 79, 104419. [Google Scholar] [CrossRef] [Scilit]
- Pauncz, R. The Construction of Spin Eigenfunctions: An Exercise Book; Springer Science & Business Media: Berlin/Heidelberg, Germany, 2012. [Google Scholar]
- Sahoo, S.; Rajamani, R.; Ramasesha, S.; Sen, D. Fully symmetrized valence-bond based technique for solving exchange Hamiltonians of molecular magnets. Phys. Rev. B 2008, 78, 054408. [Google Scholar]
- Sahoo, S.; Ramasesha, S. Full spin and spatial symmetry adapted technique for correlated electronic hamiltonians: Application to an icosahedral cluster. Int. J. Quantum Chem. 2012, 112, 1041–1054. [Google Scholar] [CrossRef] [Scilit]
- Bernu, B.; Lecheminant, P.; Lhuillier, C.; Pierre, L. Exact spectra, spin susceptibilities, and order parameter of the quantum Heisenberg antiferromagnet on the triangular lattice. Phys. Rev. B 1994, 50, 10048. [Google Scholar] [CrossRef] [Scilit]
- Löwdin, P.-O. Quantum Theory of Many-Particle Systems. III. Extension of the Hartree-Fock Scheme to Include Degenerate Systems and Correlation Effects. Phys. Rev. 1955, 97, 1509. [Google Scholar]
- Ghassemi Tabrizi, S.; Kühne, T.D. Analytical Solutions of Symmetric Isotropic Spin Clusters Using Spin and Point Group Projectors. Magnetism 2024, 4, 183–199. [Google Scholar] [CrossRef] [Scilit]
- Ghassemi Tabrizi, S.; Kühne, T.D. Projective Spin Adaptation for the Exact Diagonalization of Isotropic Spin Clusters. Magnetism 2024, 4, 332–347. [Google Scholar] [CrossRef] [Scilit]
- Altmann, S.L.; Herzig, P. Point-Group Theory Tables; Oxford University Press: Oxford, UK, 1994. [Google Scholar]
- Percus, J.K.; Rotenberg, A. Exact Eigenfunctions of Angular Momentum by Rotational Projection. J. Math. Phys. 1962, 3, 928–932. [Google Scholar] [CrossRef] [Scilit]
- Pauncz, R. Spin Eigenfunctions: Construction and Use; Springer Science & Business Media: Berlin/Heidelberg, Germany, 1979. [Google Scholar]
- Smith, V.H., Jr.; Harris, F.E. Projection of Exact Spin Eigenfunctions. J. Math. Phys. 1969, 10, 771–778. [Google Scholar] [CrossRef] [Scilit]
- Harbrecht, H.; Peters, M.; Schneider, R. On the low-rank approximation by the pivoted Cholesky decomposition. Appl. Numer. Math. 2012, 62, 428–440. [Google Scholar] [CrossRef] [Scilit]
- Lehtola, S. Curing basis set overcompleteness with pivoted Cholesky decompositions. J. Chem. Phys. 2019, 151, 241102. [Google Scholar] [CrossRef] [Scilit]
- Konstantinidis, N.P. Antiferromagnetic Heisenberg model on the icosahedron: Influence of connectivity and the transition from the classical to the quantum limit. J. Phys. Condens. Matter 2015, 27, 076001. [Google Scholar] [CrossRef] [Scilit]
- Borrás-Almenar, J.J.; Clemente-Juan, J.M.; Coronado, E.; Tsukerblat, B.S. MAGPACK1 A package to calculate the energy levels, bulk magnetic properties, and inelastic neutron scattering spectra of high nuclearity spin clusters. J. Comput. Chem. 2001, 22, 985–991. [Google Scholar] [CrossRef] [Scilit]
- Schnack, J. (Universität Bielefeld, Bielefeld, Germany). Personal communication, 2025.
Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2025 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license (https://creativecommons.org/licenses/by/4.0/).