1. Introduction
According to the Intergovernmental Panel on Climate Change (IPCC) Sixth Assessment Report [
1], the industrial sector remains one of the most challenging areas for climate change mitigation because it combines high energy demand with process-related greenhouse gas emissions that cannot be eliminated through energy efficiency measures alone. The production of basic materials—including iron, steel and non-ferrous metals—represents the largest share of industrial emissions due to the need for high-temperature processing and the continued reliance on fossil fuels. The report highlights that achieving climate neutrality will require a combination of low-carbon electricity, process electrification, hydrogen-based technologies, improved material efficiency, circular resource use and carbon capture where direct emissions cannot be avoided. Rather than relying on a single technological solution, the transition of the metallurgical industry will depend on the integration of multiple complementary strategies capable of reducing both energy consumption and process-related emissions while maintaining industrial productivity and competitiveness.
The decarbonisation of high-temperature metallurgical operations is no longer treated as a single-fuel substitution problem. The recent literature increasingly presents the transition as a combined process-engineering challenge in which the heat source, the reducing agent, the reactor atmosphere and the solids’ handling concept must be redesigned together. This point is particularly important for rotary and moving-bed systems, where the material is continuously mixed, the gas phase is in direct contact with the reacting solids and the local oxygen potential can change rapidly along the reactor length. In this context, electrification and hydrogen use are not independent solutions; their real value appears when they are integrated into reactor architectures that can control temperature and gas composition at the same time.
Recent work on industrial process heat has shown that electrification and hydrogen will both be required for deep decarbonisation, but their suitability depends strongly on temperature level, process configuration and the role of the gas phase. Leicher et al. [
2] note that electrical heating can provide high controllability and direct coupling with renewable power, while hydrogen can replace fossil fuels in processes where a gaseous fuel or reducing agent is required. For metallurgical processing, this distinction is essential. Electrical heating can remove combustion products from the reactor, but it does not automatically create a stable reducing atmosphere. Process-level assessments of electrified mineral processing similarly indicate that its environmental benefit depends on the electricity mix, thermal integration and configuration of the electrically heated reactor [
3]. Conversely, hydrogen can provide a carbon-free reduction route, but only if the reactor is sufficiently tight and well controlled to avoid air ingress, hydrogen dilution and unnecessary gas losses.
This has direct relevance to extractive metallurgy. The steel sector has been the main reference point for hydrogen-based process redesign because it combines very high energy demand with direct carbon use in reduction reactions. Recent MDPI studies have examined how hydrogen direct reduction, electrification and changes in energy structure can be combined to reduce emissions from steelmaking [
4,
5]. These studies are useful beyond ironmaking because they show that the low-carbon transition depends on the coupling between renewable electricity, hydrogen supply, heat generation and process control. In practice, this means that a new reactor cannot be assessed only by its energy source—it must also be evaluated by its ability to maintain a useful reaction environment under realistic operating conditions.
The hydrogen reduction of iron oxides has been widely reviewed in the recent literature. Zakeri et al. [
6] emphasised that the reduction behaviour is affected not only by temperature and gas composition, but also by impurities and phase evolution within the solid. Heidari et al. [
7] similarly showed that hydrogen-reduction kinetics depend on ore type, porosity, particle size, temperature and gas-flow conditions. These findings reinforce a central point for rotary reactors: the gas–solid contact is beneficial only when it occurs under a controlled atmosphere. If oxygen ingress or water-vapour accumulation is not managed, the effective H
2/H
2O ratio decreases and the reduction potential of the gas phase is weakened.
More recent studies have moved from general feasibility towards reactor-scale behaviour and process optimisation. Ji et al. [
8] reviewed the development and application of hydrogen-based direct reduction and highlighted the need for improved integration between reduction reactors, hydrogen supply and downstream processing. Experimental and modelling work by Cavaliere et al. [
9] on the hydrogen reduction of pellets confirmed that temperature, exposure time and gas transport strongly influence the degree of reduction. Özgün et al. [
10] further showed that hydrogen pressure can modify reduction kinetics and microstructural evolution. Although these studies mainly concern iron ore pellets, they provide an important mechanistic basis for the hydrogen-assisted treatment of more complex oxide feedstocks, including nickel-bearing laterites.
The extension of hydrogen metallurgy to non-ferrous and complex ores is now receiving increasing attention. Nickel laterites are especially relevant because they are abundant, chemically heterogeneous and increasingly important for battery and stainless-steel supply chains. Conventional laterite processing relies on energy-intensive pyrometallurgical routes or acid-based hydrometallurgical flowsheets. Selective reduction studies further show that the reduction temperature, reductant availability and mineralogical association of nickel and iron determine ferronickel formation and subsequent metal recovery [
11]. Kinetic analysis of laterite calcination has shown that several overlapping transformations occur below 1000 °C, including dehydroxylation, carbonate decomposition and structural changes that subsequently influence oxide reducibility [
12]. Recent work has therefore explored hydrogen-assisted routes as a way to reduce direct carbon use and improve selectivity [
13]. Wijenayake et al. [
13] investigated ferronickel production from limonitic laterite using hydrogen reduction combined with cementation, demonstrating that hydrogen can participate effectively in laterite processing. Fan et al. [
14] reviewed the comprehensive utilisation of nickel laterite ores and identified hydrogen and other clean reductants as promising options for future low-carbon flowsheets.
A particularly important recent development is the work of Manzoor et al. [
15], who demonstrated a hydrogen-based reduction of laterites for sustainable nickel production. Their study is significant because it shows that hydrogen routes are not limited to high-grade or simple oxide feedstocks. Instead, hydrogen can be used to target complex mineral assemblages when the process conditions are properly designed. Investigations employing gaseous reductants have also confirmed that temperature and exposure time control the sequential transformation of iron oxides and the formation and growth of Fe–Ni metallic phases [
16]. Earlier hydrogen-reduction experiments on nickel laterite also showed that additives such as sodium sulphate can promote nickel liberation and ferronickel formation, although relatively long treatment periods were required [
17]. For an electrified rotary system, this literature supports the idea that laterite pre-reduction can become faster and cleaner, provided that temperature, residence time and gas composition are held within a narrow and reproducible operating window.
The importance of gas-atmosphere control is also evident in rotary furnace studies. Chung et al. [
18] investigated the hydrogen reduction of tellurium oxide in a rotary kiln furnace and showed that controlled hydrogen treatment can promote oxide reduction under rotary processing conditions. Although the material system differs from laterite, the reactor implications are directly relevant. Rotary motion improves contact between particles and the gas phase, but it also increases the importance of sealing, gas distribution and consistent residence time. These are precisely the aspects that must be addressed if hydrogen is to be used efficiently in rotary metallurgical systems.
Hydrogen plasma studies point in the same direction from a different technological angle. Satritama et al. [
19] reviewed hydrogen plasma for low-carbon extractive metallurgy and identified both the potential of hydrogen-based reduction and the limitations associated with scale-up, heat utilisation, reoxidation and process stability. The review is useful for the present work because it confirms that hydrogen-based metallurgy is not only a question of thermodynamic feasibility. Reactor design, heat-transfer efficiency and atmosphere integrity determine whether the reducing potential of hydrogen can be translated into reproducible metallurgical performance.
In parallel, the decarbonisation of calcination processes has become a major research topic. Calcination is difficult to decarbonise because CO
2 is released not only from fuel combustion but also from carbonate decomposition. Electrification can eliminate the combustion-related part of the emissions, while airtight or indirectly heated operation can avoid dilution of the process CO
2 stream. Electrified calciner configurations can therefore simultaneously reduce combustion-related emissions and generate a more concentrated CO
2 stream that is better suited to capture and utilisation [
20]. This is important for magnesite and limestone calcination, where a concentrated CO
2 outlet stream can simplify capture, utilisation or further processing. Magnesite calcination is the principal industrial route for MgO production, while the calcination temperature and residence time strongly influence the reactivity, surface area and final properties of the produced magnesia [
21]. Recent analysis of the magnesia sector has also demonstrated that fuel substitution, process optimisation and improved kiln-energy management can substantially influence both production costs and CO
2 emissions [
22]. Ryan et al. [
23] modelled calcination in a rotary lime kiln and showed that heat transfer, reaction kinetics and particle behaviour must be considered together. Zhu et al. [
24] also demonstrated through modelling that limestone decomposition is governed by coupled heat transfer, mass transfer and chemical reaction steps.
Recent MDPI work on carbonate decomposition provides additional support for this interpretation. Zhuang et al. [
25] studied the thermal decomposition of calcium carbonate under different atmospheres and heating rates, showing that decomposition temperature and reaction rate are strongly linked to heating conditions and atmosphere. While calcium carbonate differs from magnesite, the underlying carbonate-decomposition logic is comparable: reaction progress depends on the ability of the reactor to supply heat uniformly and remove the evolved gas without creating local mass-transfer limitations. For magnesite calcination in an electrified indirect rotary kiln, this means that temperature uniformity and controlled gas removal are central to achieving high conversion within a practical treatment time.
The recent literature on industrial thermal applications of hydrogen also supports the need for careful reactor design. Mirshokraee et al. [
26] reviewed hydrogen substitution for natural gas in thermal industrial applications and highlighted that hydrogen use changes combustion behaviour, safety requirements and process-control needs. Even when hydrogen is not combusted but used as a reducing gas, the same broader conclusion remains valid: hydrogen-compatible equipment must be designed around gas tightness, monitoring, ventilation and operating stability. Joyo et al. [
27] similarly showed that hydrogen integration in steel decarbonisation requires a techno-economic evaluation of supply, process efficiency and operational constraints.
Taken together, the most recent MDPI and Scopus-indexed literature shows that the next generation of low-carbon metallurgical reactors must move beyond simple electrification. Electrically heated systems offer clear advantages by removing combustion gases and improving thermal control. Experimental investigation of an externally heated rotary calciner has demonstrated the technical feasibility of transferring heat through the kiln wall, with radiation providing the dominant heat-transfer contribution at calcination temperatures [
28]. Hydrogen processing requires an additional layer of atmosphere management. Rotary systems are attractive because they provide solids mixing, particle renewal and flexible residence time; however, conventional direct-fired rotary kilns are poorly suited to hydrogen operation because they do not fully isolate the heat source from the process atmosphere. This creates a clear technological gap for sealed, indirectly heated rotary reactors able to control temperature and gas composition independently.
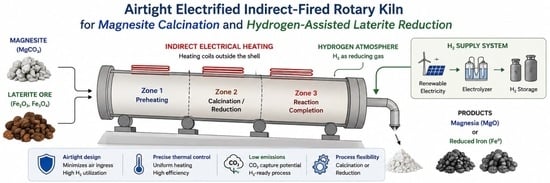
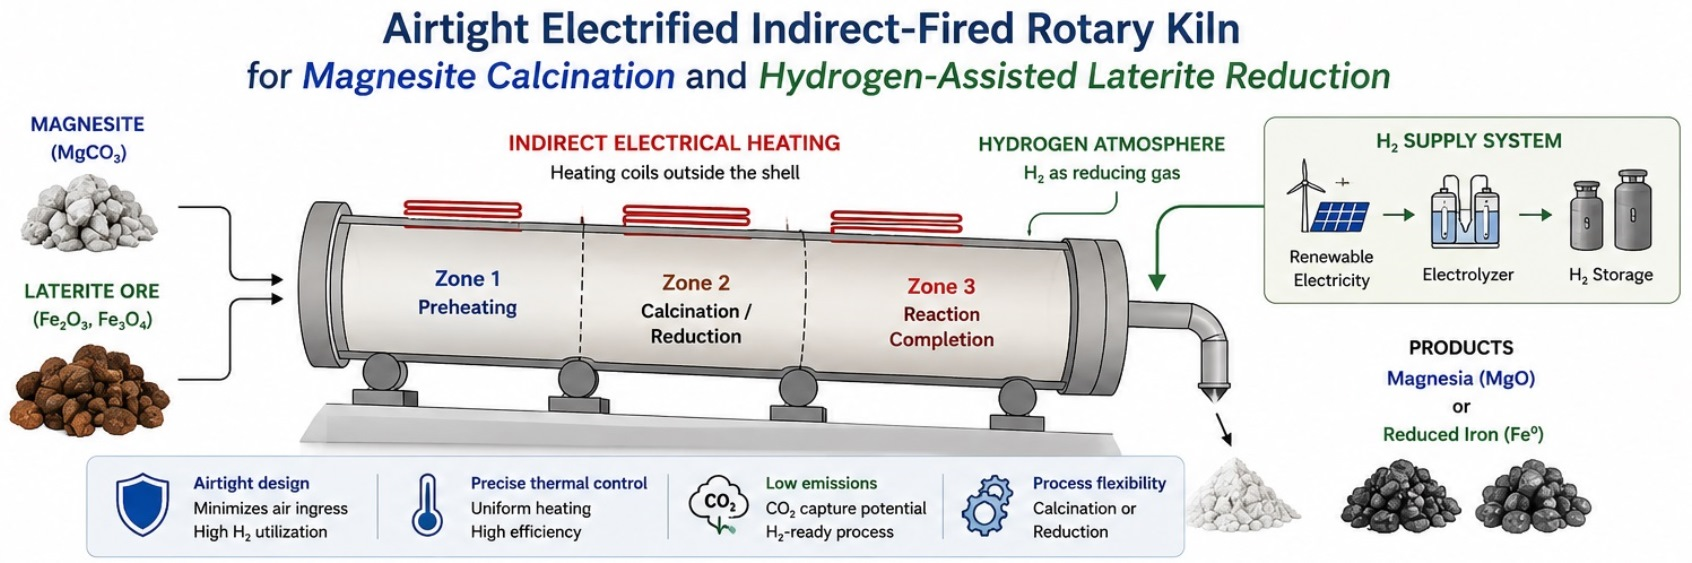
In this framework, this study presents the engineering of a fully sealed reaction chamber assembled in an indirect electrically heated rotary kiln to investigate how reactor electrification and atmosphere isolation can be combined within a single processing unit. Rather than focusing solely on replacing conventional burners with electrical heating, the reactor was engineered to provide independent control of heat transfer and gas composition, creating stable conditions for thermochemical reactions while reducing air ingress, thermal losses and unnecessary consumption of reducing gases. To examine its performance under representative metallurgical conditions, the system was tested in two different applications: magnesite calcination and hydrogen-assisted reduction of lateritic ores. The behaviour of both the feed and treated materials was evaluated through physicochemical characterisation, whereas temperature profile and process gas composition were monitored continuously throughout each experimental campaign. The results provide an assessment of the reactor’s ability to maintain accurate processing conditions during high-temperature treatment and demonstrate its potential as a flexible process for future electrified and hydrogen-compatible metallurgical applications.
The airtight electrified indirect-fired rotary kiln examined in the present study is positioned within this gap. Its relevance lies in the integration of three elements that are often treated separately in the literature: electrical heat supply, atmosphere isolation and hydrogen-compatible rotary processing. By separating the heat source from the reaction chamber, the system avoids combustion-gas dilution and allows the gas phase to be selected according to the process requirement. During magnesite calcination, this can support controlled CO
2 evolution and concentrated off-gas formation. During laterite treatment, it can preserve a reducing hydrogen atmosphere and promote oxide reduction under reproducible conditions. “The novelty of the proposed reactor does not lie simply in replacing conventional fuels with electricity or hydrogen, but in integrating indirect electrical heating, atmosphere isolation and hydrogen-compatible rotary processing within a single reactor architecture specifically designed for high-temperature metallurgical applications.” The materials, experimental procedures and analytical methods adopted in this work are described in the
Section 2.
2. Materials and Methods
2.1. Integrated Methodological Framework
Two representative mineral feedstocks were selected to evaluate the applicability of the electrified indirect-fired rotary kiln under different thermochemical processing conditions. A cryptocrystalline magnesite was used for calcination experiments, whereas a nickel-bearing lateritic ore served as the feed material for the hydrogen-reduction tests. These materials were selected because they represent two important extractive processes with fundamentally different reaction mechanisms: thermal decomposition in the case of magnesite and the solid-state reduction of metal oxides in the case of laterite.
2.2. Raw Materials Characterisation
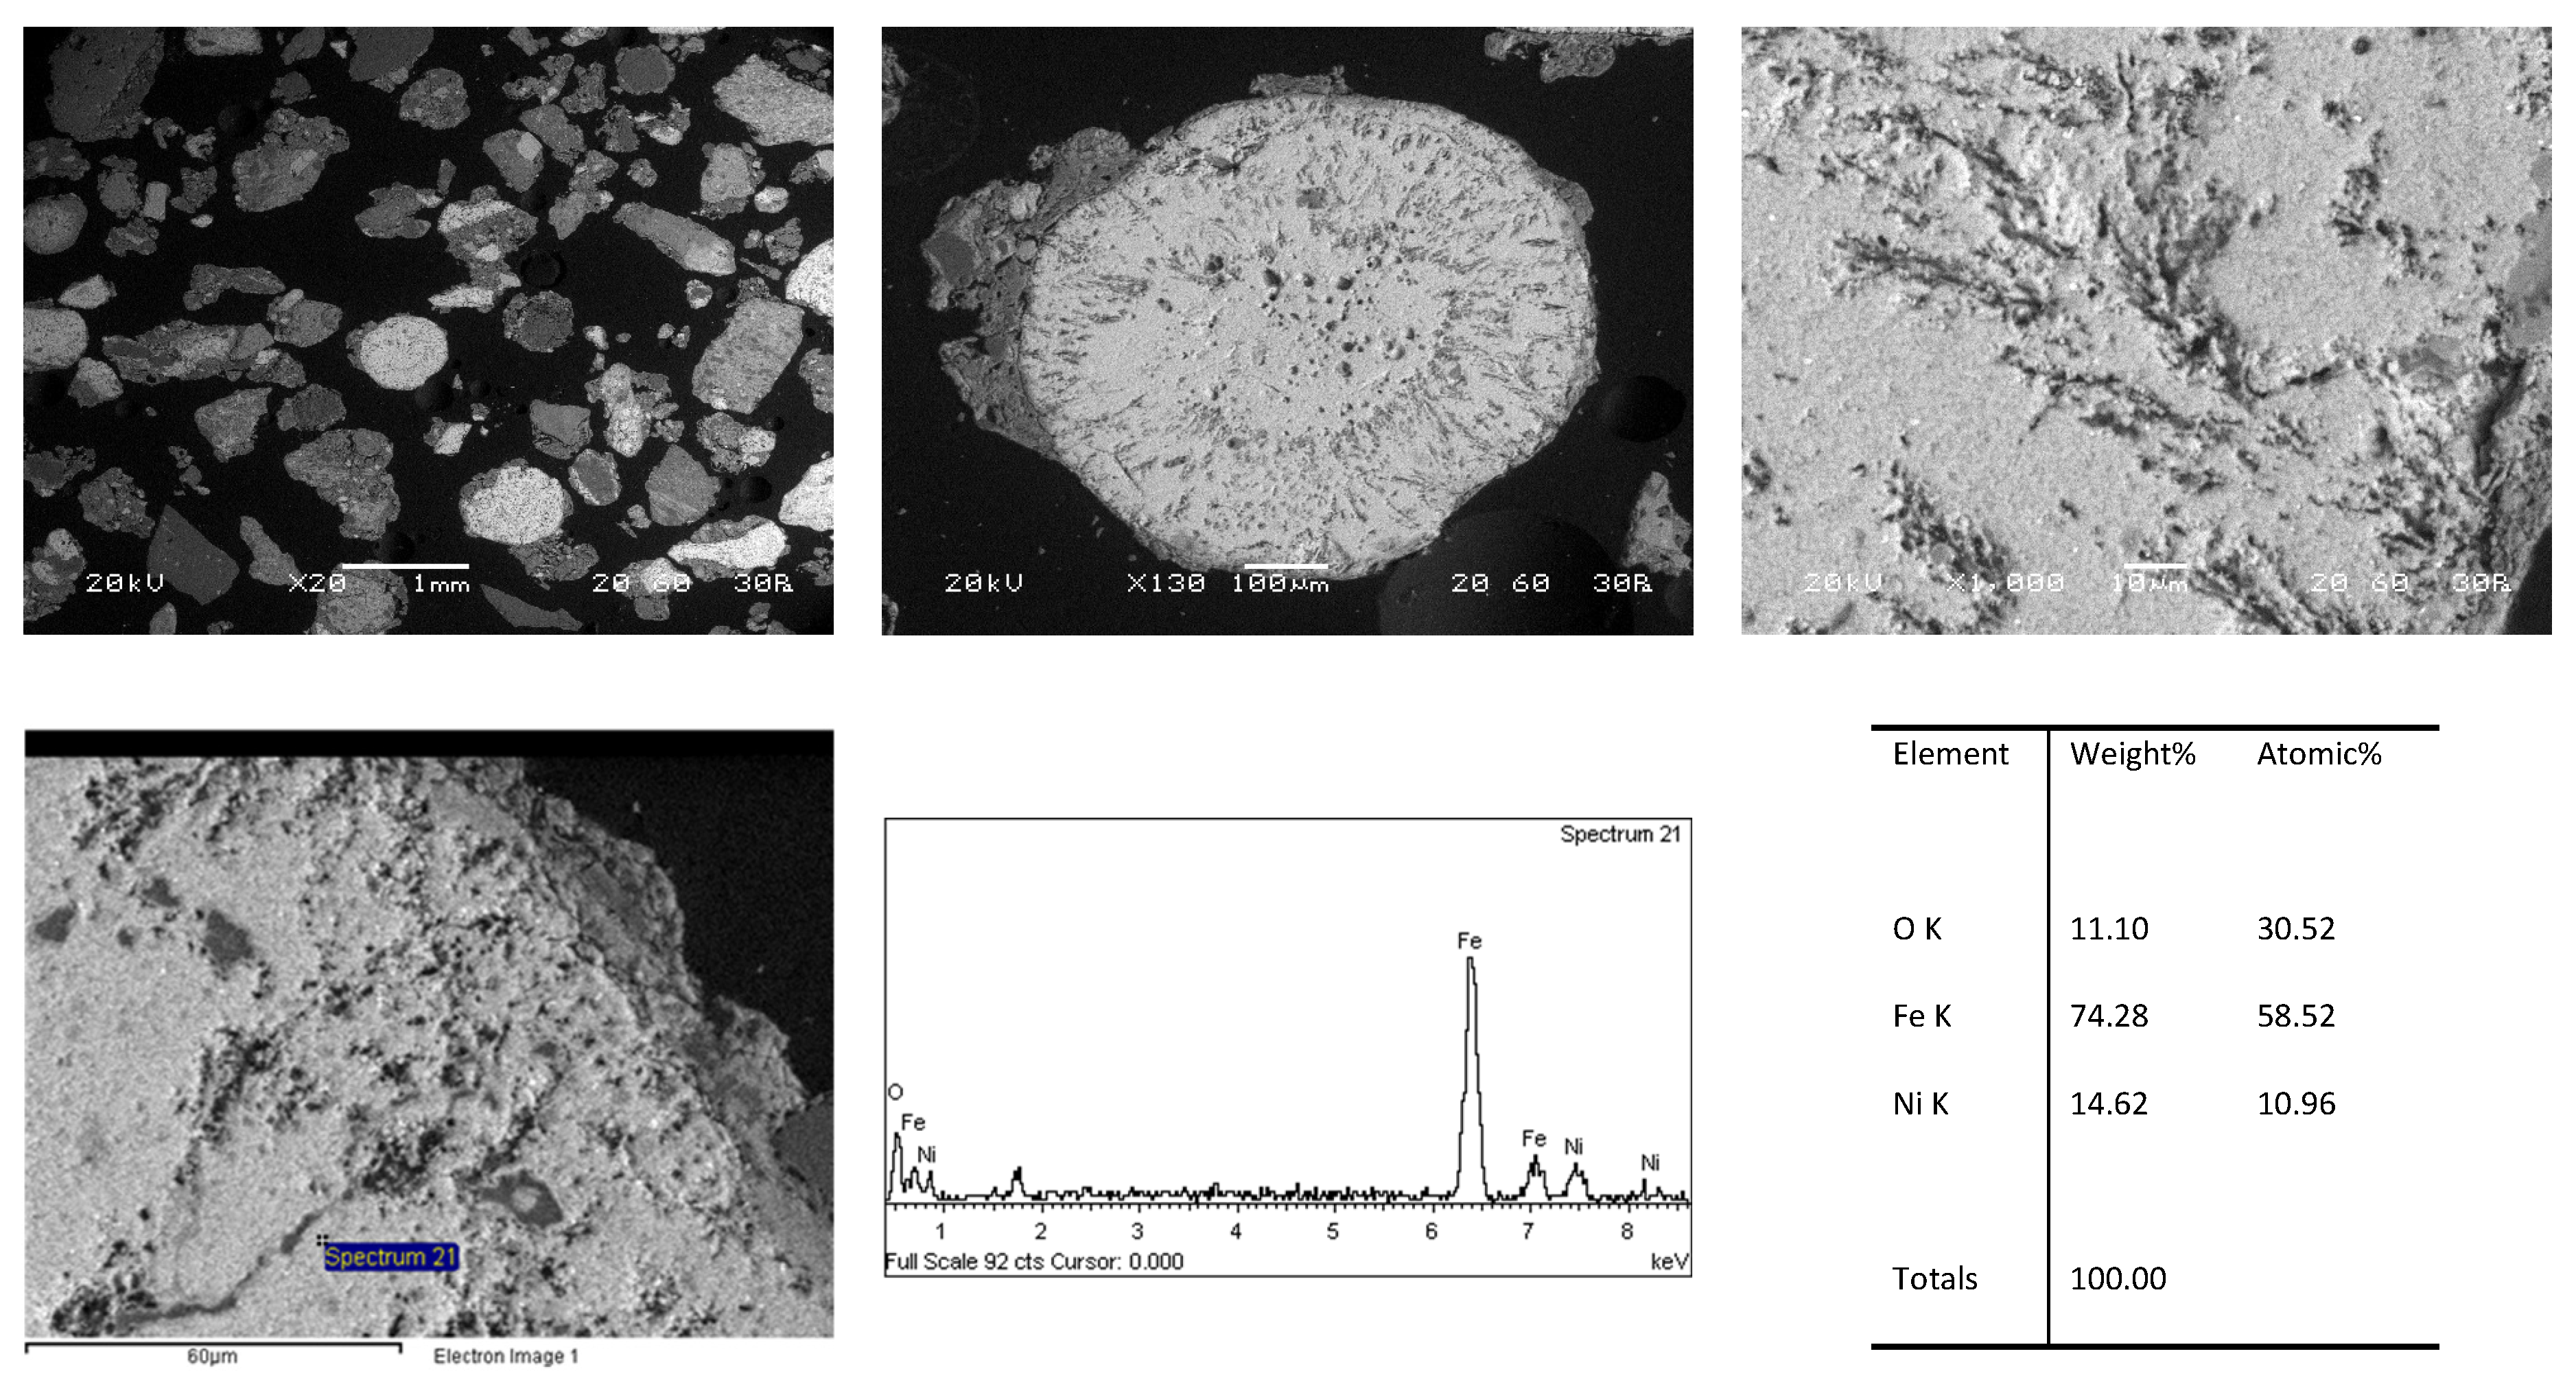
Before each experimental campaign, the feedstocks were subjected to comprehensive physicochemical characterisation to establish their initial properties and provide a reliable basis for interpreting the transformations occurring during thermal treatment. Chemical composition was determined by X-ray fluorescence (XRF), using a Spectro XEPOS (SPECTRO Analytical Instruments GmbH, Kleve, Germany) energy-dispersive spectrometer, while the mineralogical assemblage was identified by XRD employing a Bruker D8 Focus diffractometer (Bruker AXS GmbH, Karlsruhe, Germany). Particle-size distribution (PSD) measurements were carried out with a HORIBA Partica LA-960V2 laser diffraction particle-size analyser (HORIBA Ltd., Kyoto, Japan), while the coarse fraction (>150 μm) was additionally evaluated by conventional sieve analysis. For the calcination tests, thermal behaviour was investigated using simultaneous thermogravimetric and differential thermal analysis (TG-DTA) using a SETARAM THEMYS ONE thermal analyser (SETARAM–KEP Technologies, Caluire-et-Cuire, France) under an argon atmosphere. Following the laterite reduction experimental campaigns, selected samples were further examined by using a JEOL JSM-5600 SEM (JEOL Ltd., Akishima, Tokyo, Japan) equipped with an Oxford ISIS 300 EDS detector (Oxford Instruments plc, Abingdon, UK), operating at an accelerating voltage of 20 kV.
To ensure the reliability of the experimental data, all measuring instruments were operated within their specified performance limits and were calibrated before the experimental campaign. The analytical equipment used for material characterisation, including the XRF, XRD, laser diffraction particle-size analyser, TG-DTA and SEM-EDS systems, provides measurement uncertainties that are small compared with the changes observed during thermal treatment. During reactor operation, furnace temperature was monitored using a Class 1 K-type thermocouple with an accuracy of ±1.5 °C (or ±0.4% of the measured value), while the PID controller maintained the selected temperature within approximately ±1 °C of the setpoint. The hydrogen and nitrogen flow rates were regulated by a calibrated mass-flow controller with an accuracy of approximately ±1% of full scale, and the outlet-gas composition was continuously measured using a portable infrared gas analyser with a typical accuracy of ±2% of full scale. These uncertainties are significantly lower than the experimental variations observed during calcination and hydrogen reduction, providing confidence that the measured trends reflect the behaviour of the process rather than limitations of the instrumentation.
The post-treatment characterisation combined XRD and SEM–EDS analyses to obtain complementary information on the reduction process. XRD was used to identify changes in the crystalline phase assemblage, while SEM–EDS provided a comparative evaluation of the elemental composition by examining the relative changes in oxygen and metallic elements after reduction. Together, these techniques provide qualitative evidence of the progression of the reduction process and the formation of metallic phases. No quantitative metallisation measurements were performed.
2.2.1. Magnesite
This was the feed material used throughout the calcination experiments by TERNA MAG [
29] which is a leading Greek producer of high-purity magnesite and magnesia products, operating integrated mining and processing facilities in northern Euboea and supplying industrial markets worldwide. Chemical analysis indicated that the sample consisted predominantly of magnesium carbonate, with an MgCO
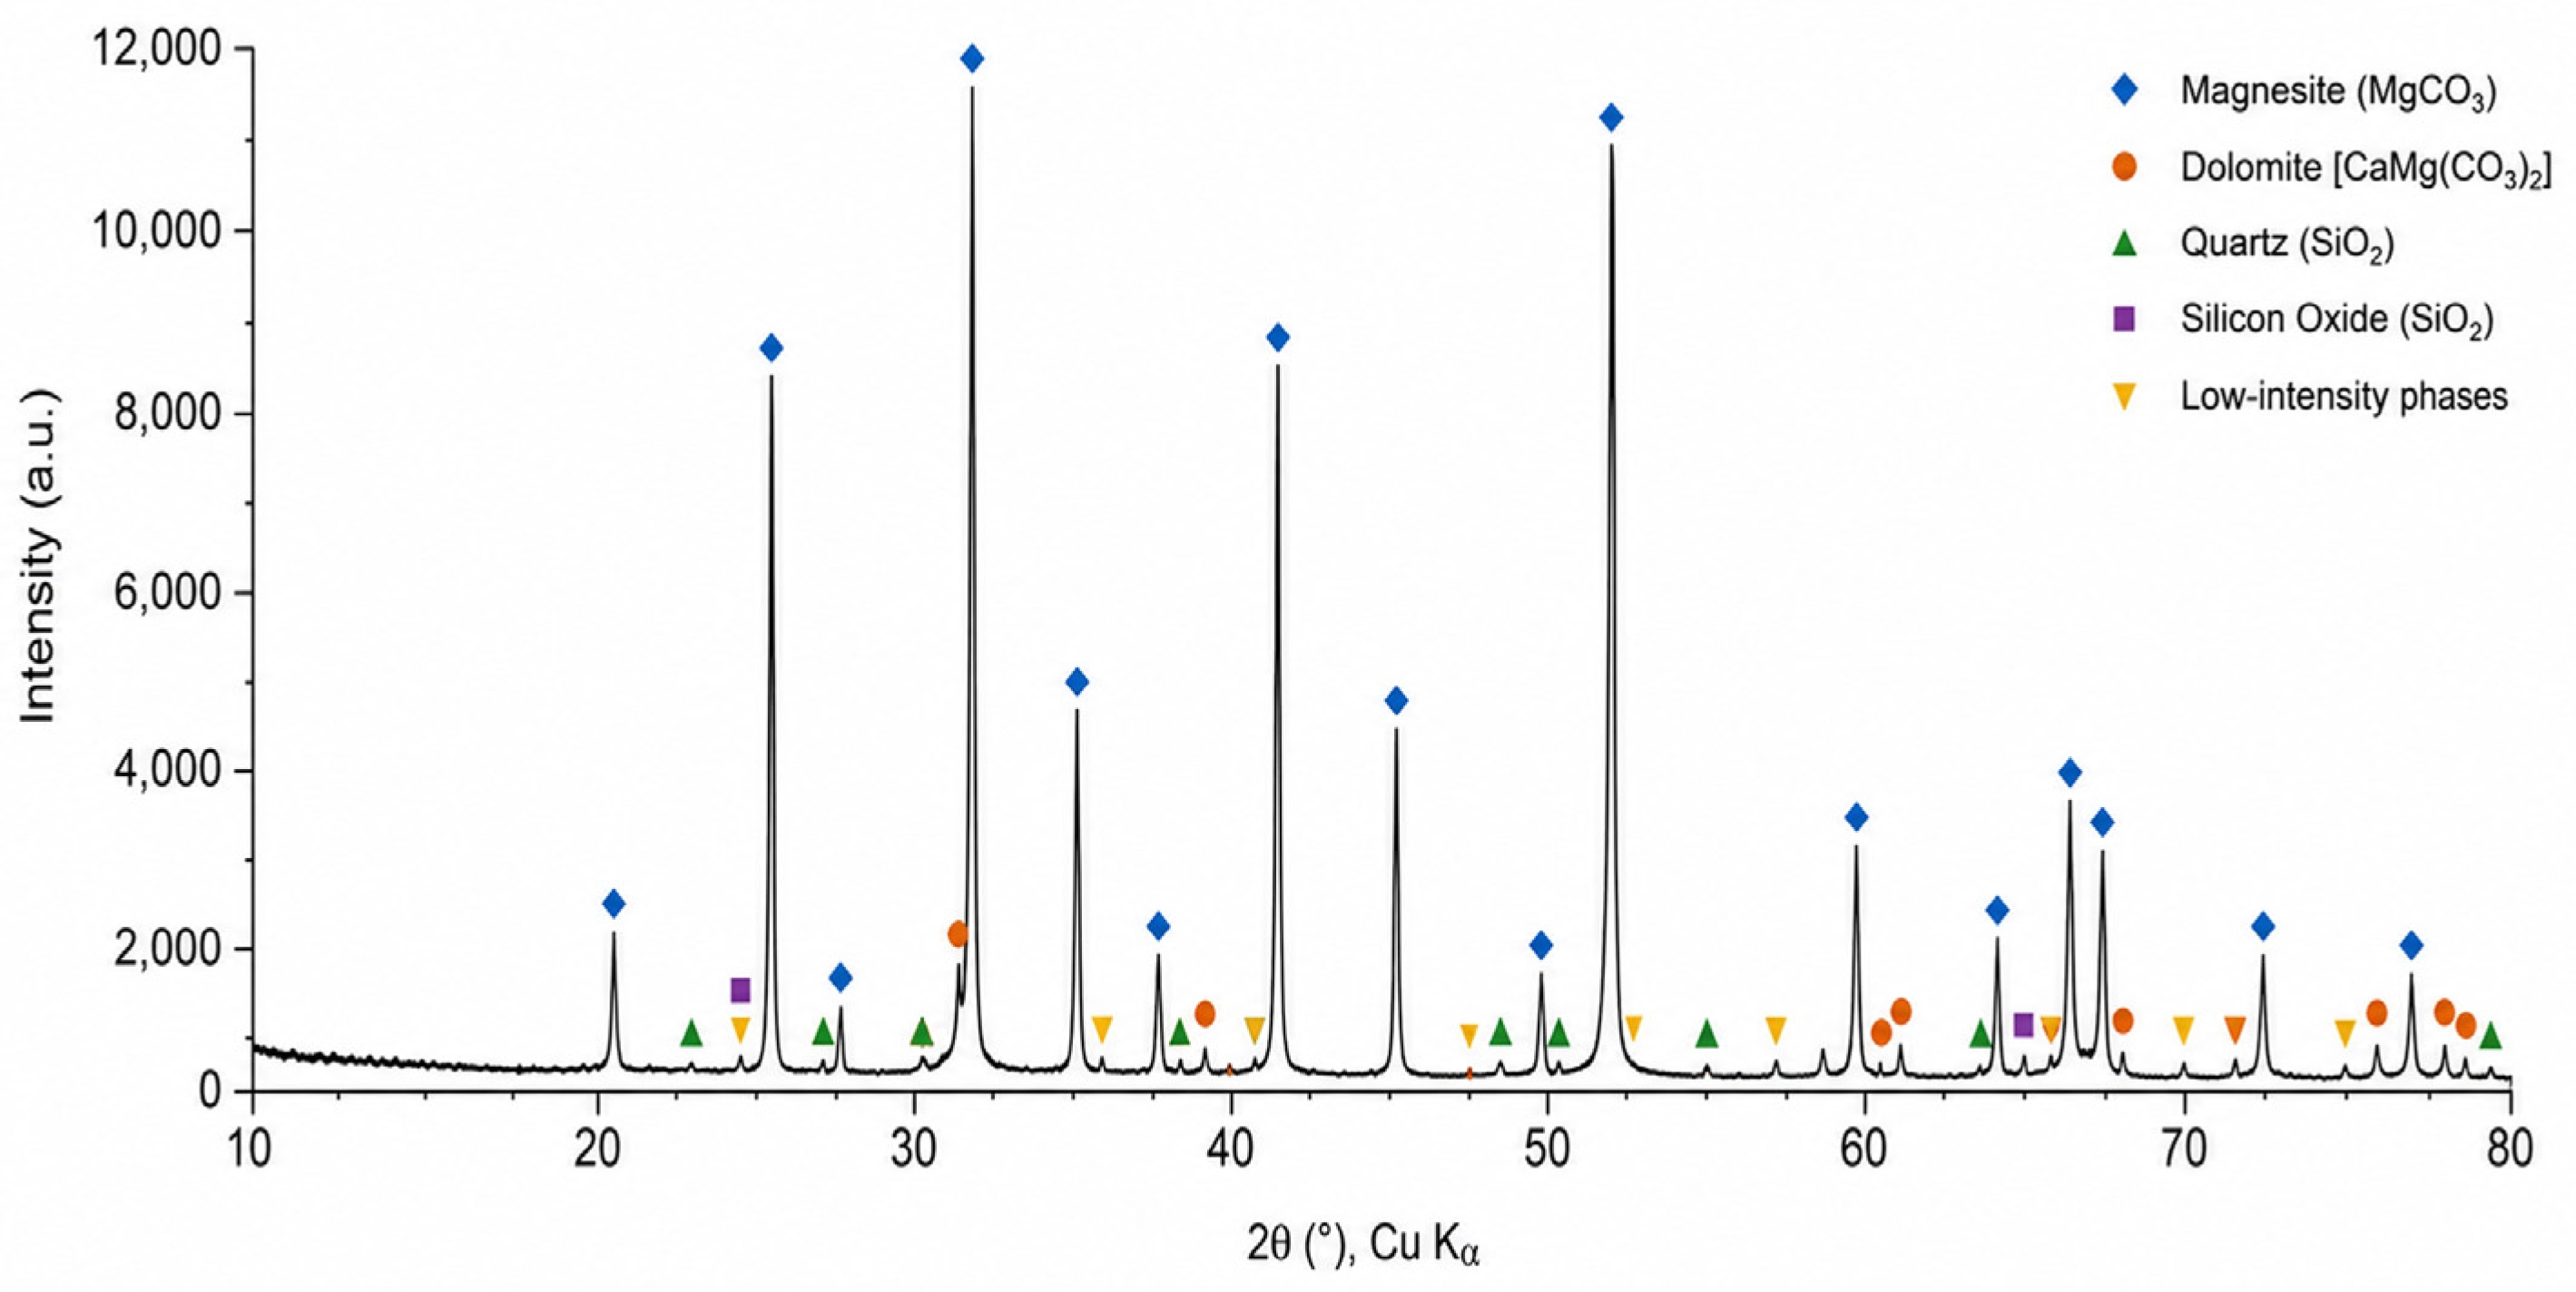
3 content of approximately 93 wt.%, confirming the high purity of the material. Minor quantities of naturally occurring gangue minerals were also present, as expected for industrial magnesite ores, but their concentration was sufficiently low that they were not expected to influence the decomposition behaviour significantly. The main mineralogical phases determined using XRD analysis in the magnesite sample, as shown in
Figure 1, were MgCO
3 (magnesite), SiO
2 (crystallised as quartz and silicon oxide) and dolomite (CaMg(CO
3)
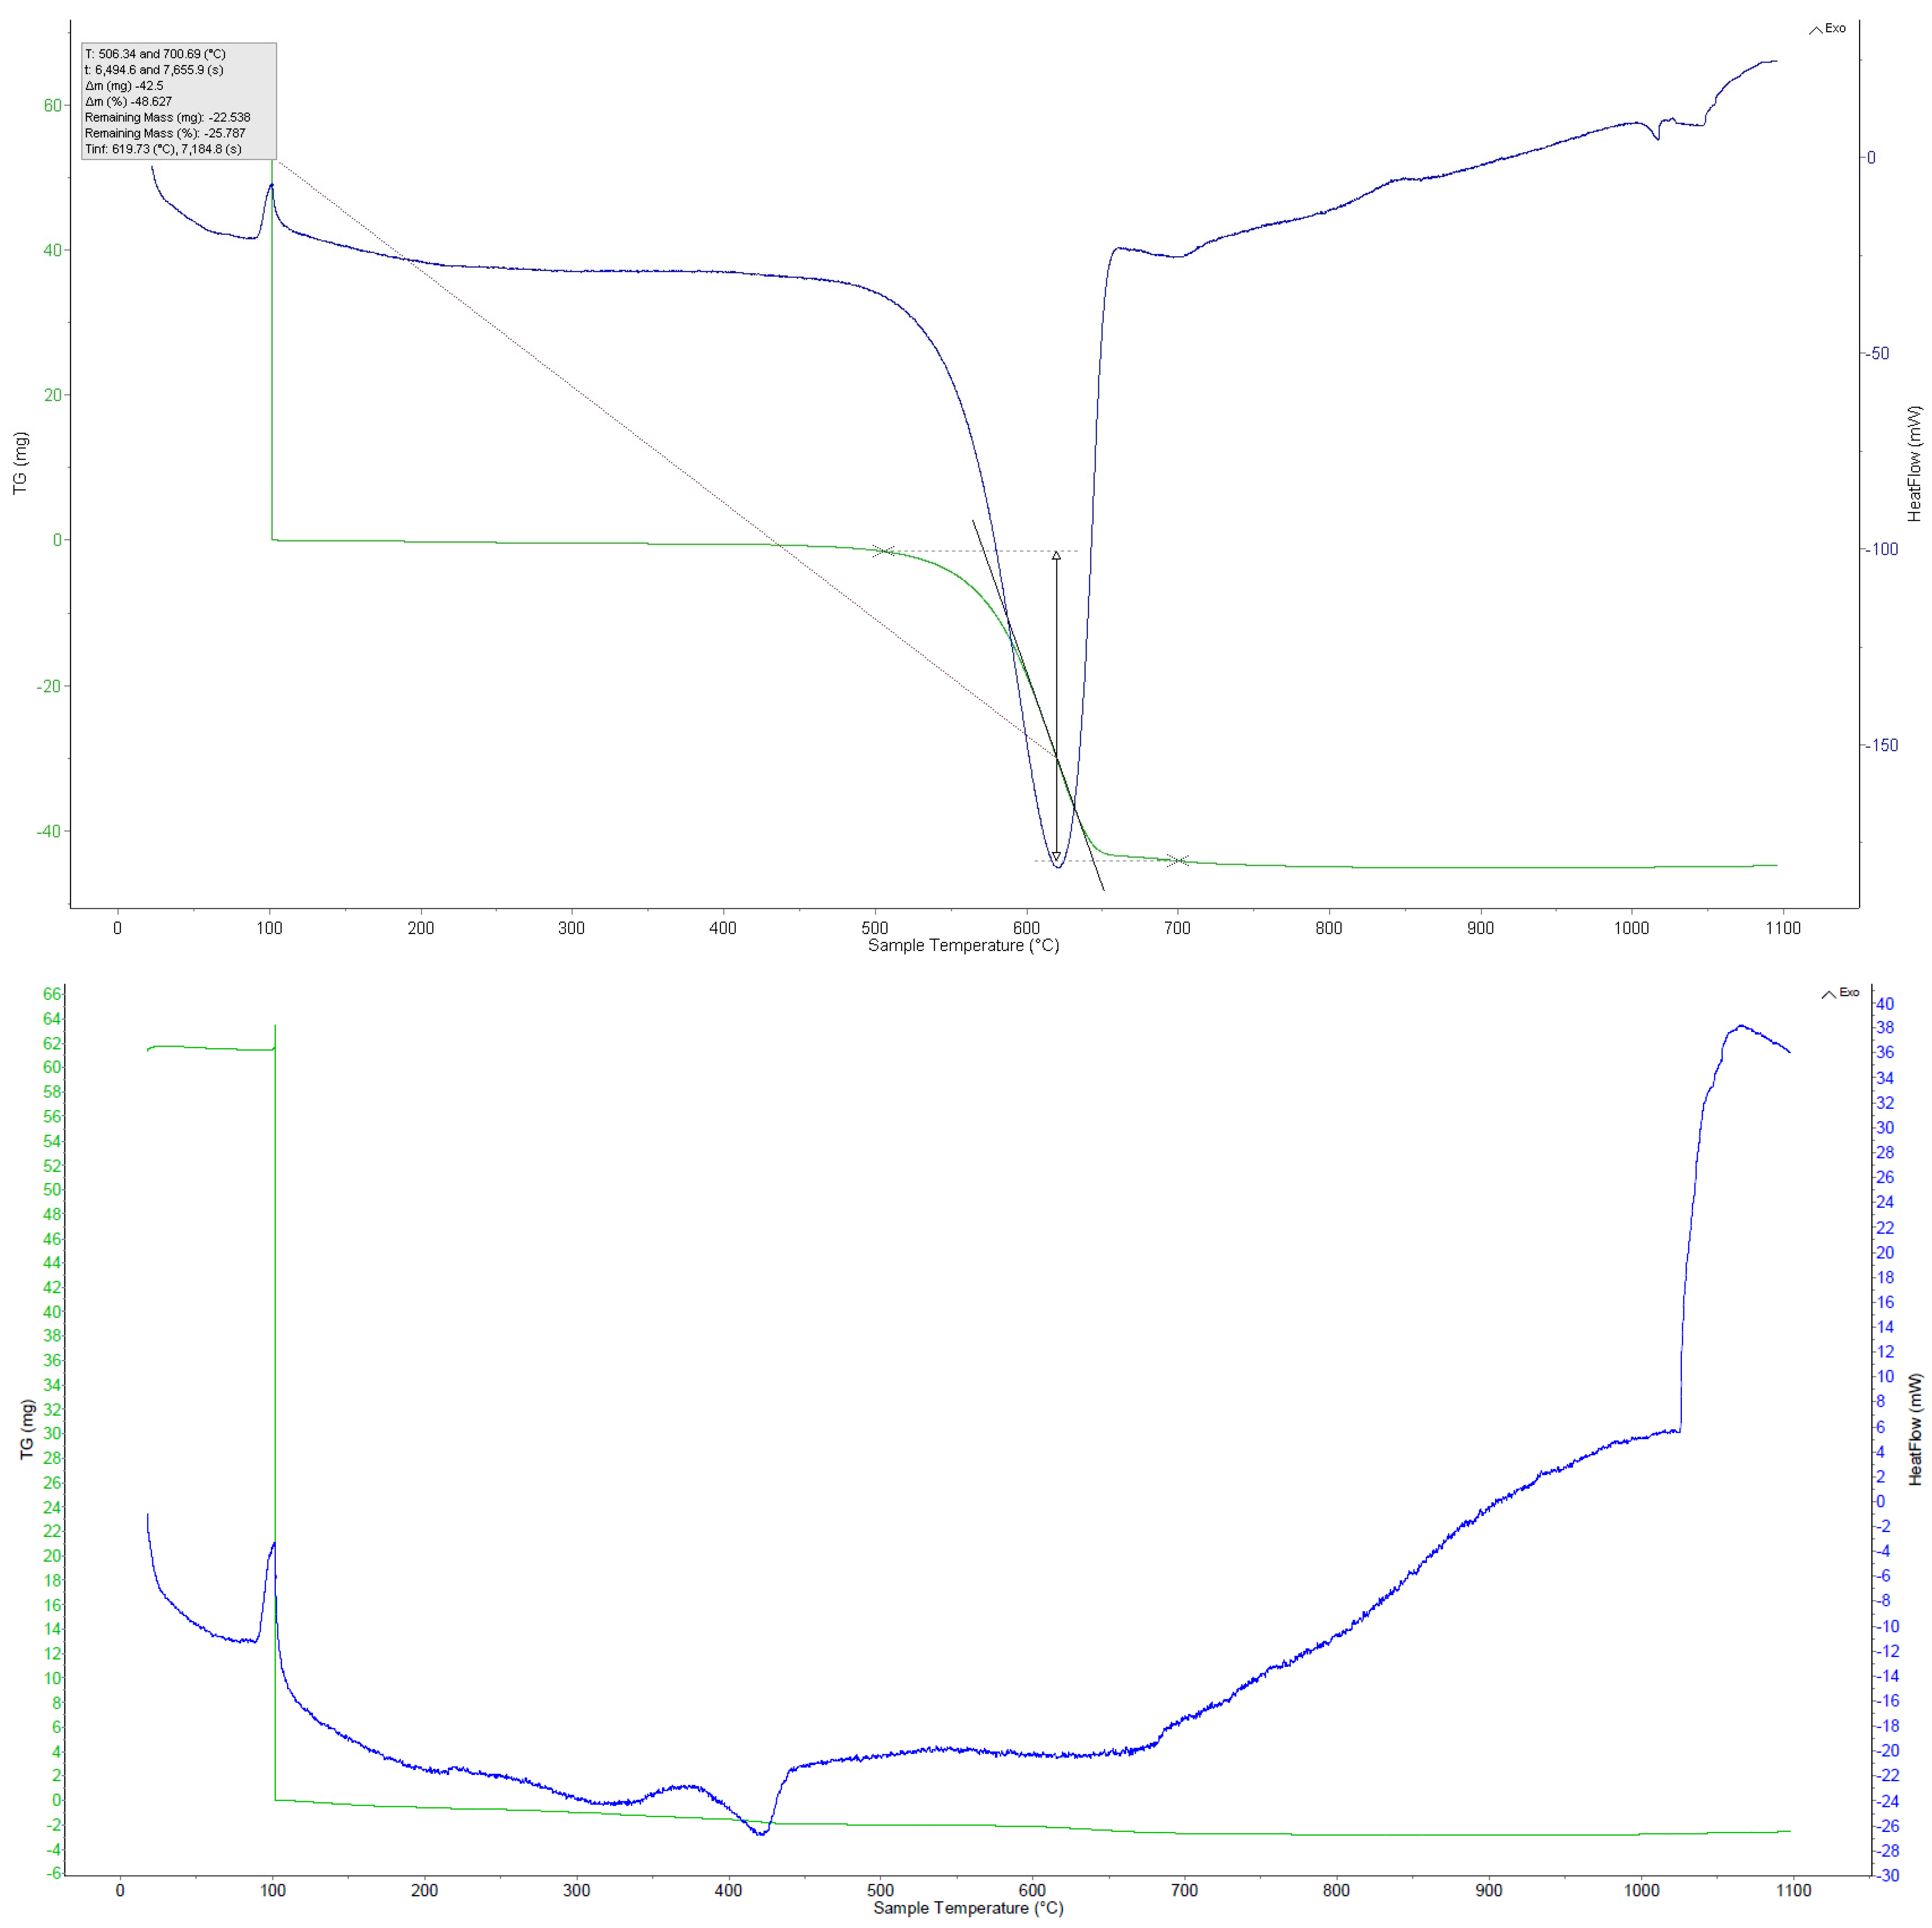
2). The thermal behaviour of the feed was assessed using simultaneous TG-DTA. The analysis revealed a single dominant decomposition stage corresponding to the decarbonation of MgCO
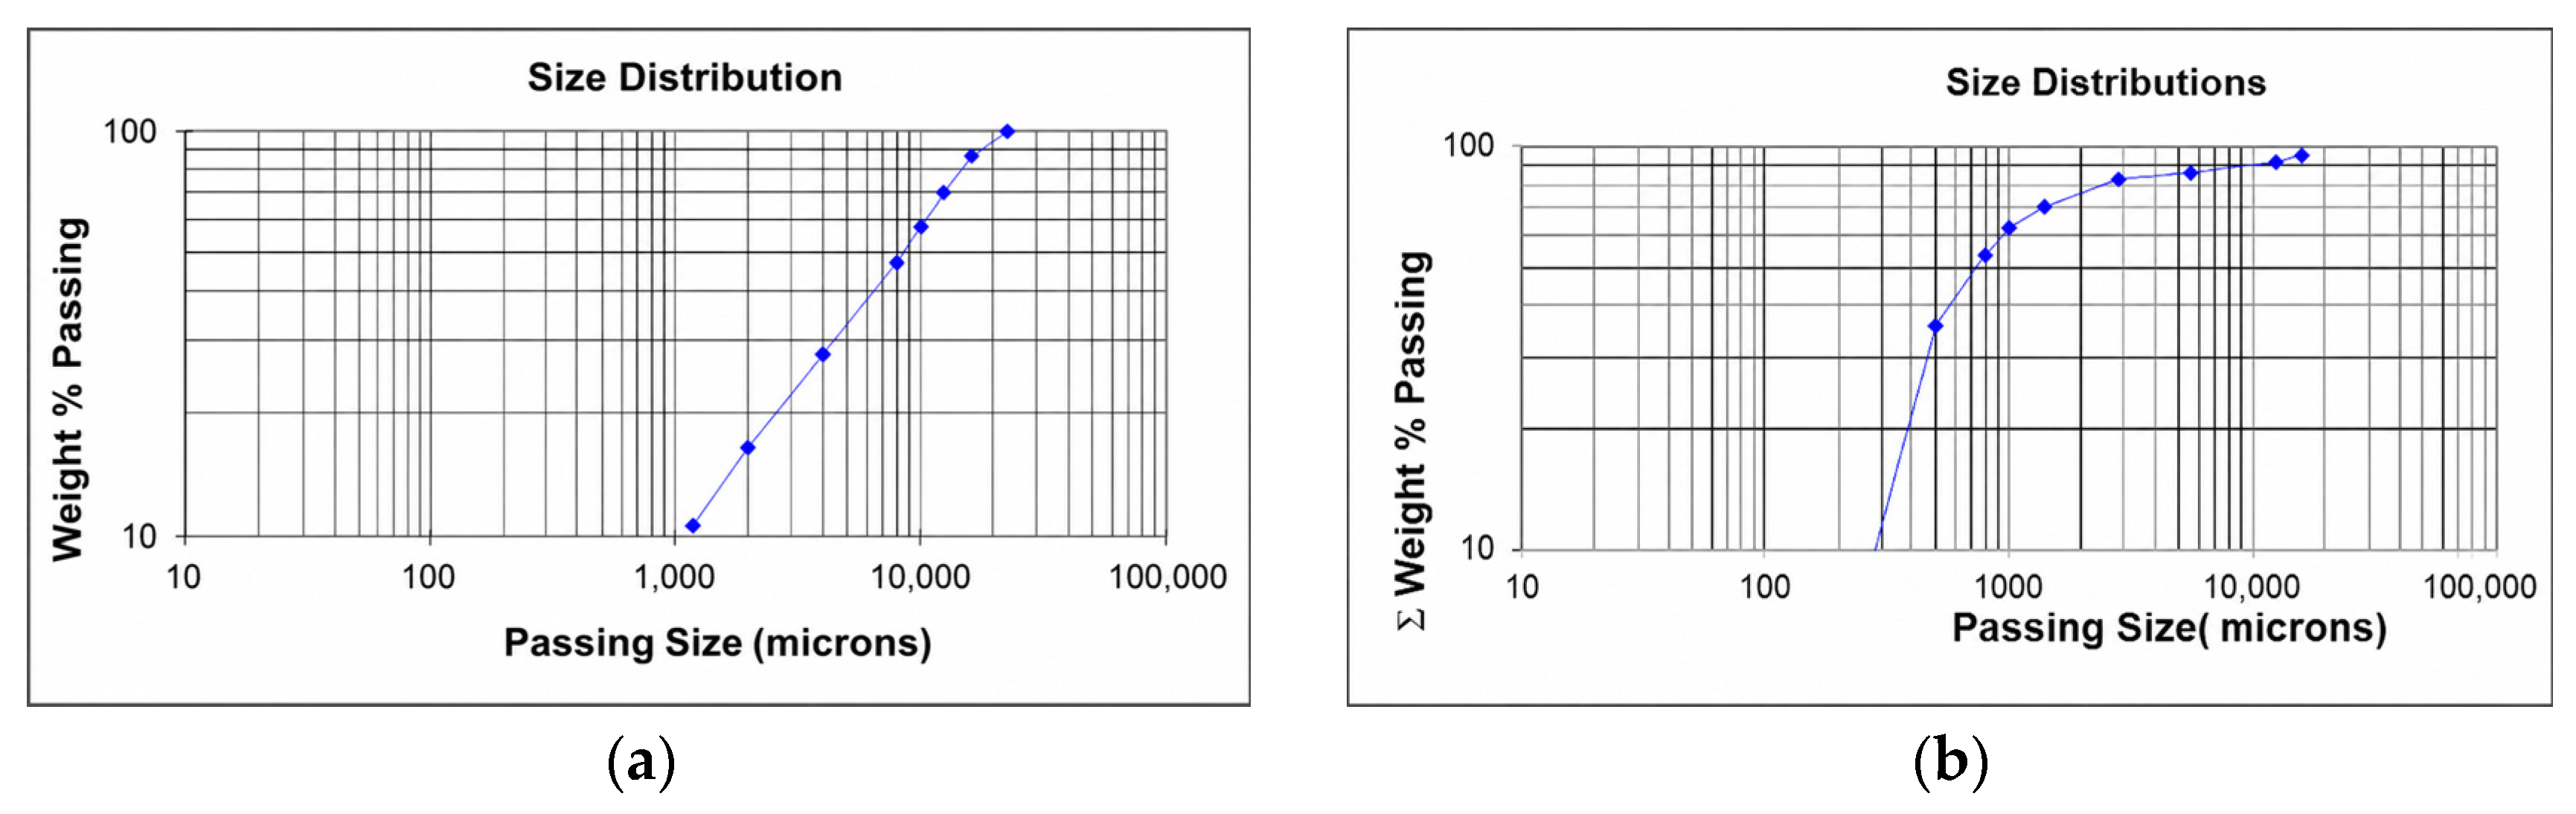
3. Weight loss began at approximately 500 °C and progressed rapidly until around 700 °C, where the decomposition reaction was essentially completed, with approximately 48.6% total mass loss. Particle-size analysis demonstrated that most particles were distributed between approximately 2 and 16 mm, although both coarser particles and a measurable fine fraction below 1.18 mm were also present. All particles were below 22.4 mm, with the majority concentrated in the intermediate size fractions; 86.7% of the sample passed the 16 mm sieve and 58.0% was finer than 10 mm. The amount of fine material was relatively limited, with 27.8%, 16.6% and 10.6% passing the 4 mm, 2 mm and 1.18 mm sieves, respectively.
2.2.2. Lateritic Ore
The lateritic ore used in this study was supplied by LARCO General Mining & Metallurgical Company S.A. [
30], one of Europe’s largest ferronickel producers, exploiting nickel-bearing laterite deposits from central Greece. Chemical composition was determined by XRF, revealing a predominantly silicate–iron oxide material. Silicon dioxide was the major constituent (41.37 wt.%), followed by Fe
2O
3 (28.57 wt.%), indicating that the ore is primarily composed of silicate minerals with abundant iron oxides. Moderate concentrations of MgO (7.51 wt.%) and Al
2O
3 (7.07 wt.%) were also detected, while CaO accounted for 3.53 wt.%. Chromium was present at relatively high levels (3.03 wt.% Cr
2O
3), reflecting the occurrence of chromium-bearing spinel minerals commonly associated with lateritic deposits. Nickel, the target metal for the reduction process, was measured at 1.05 wt.% as NiO. Minor quantities of Na
2O, K
2O, TiO
2, MnO, V
2O
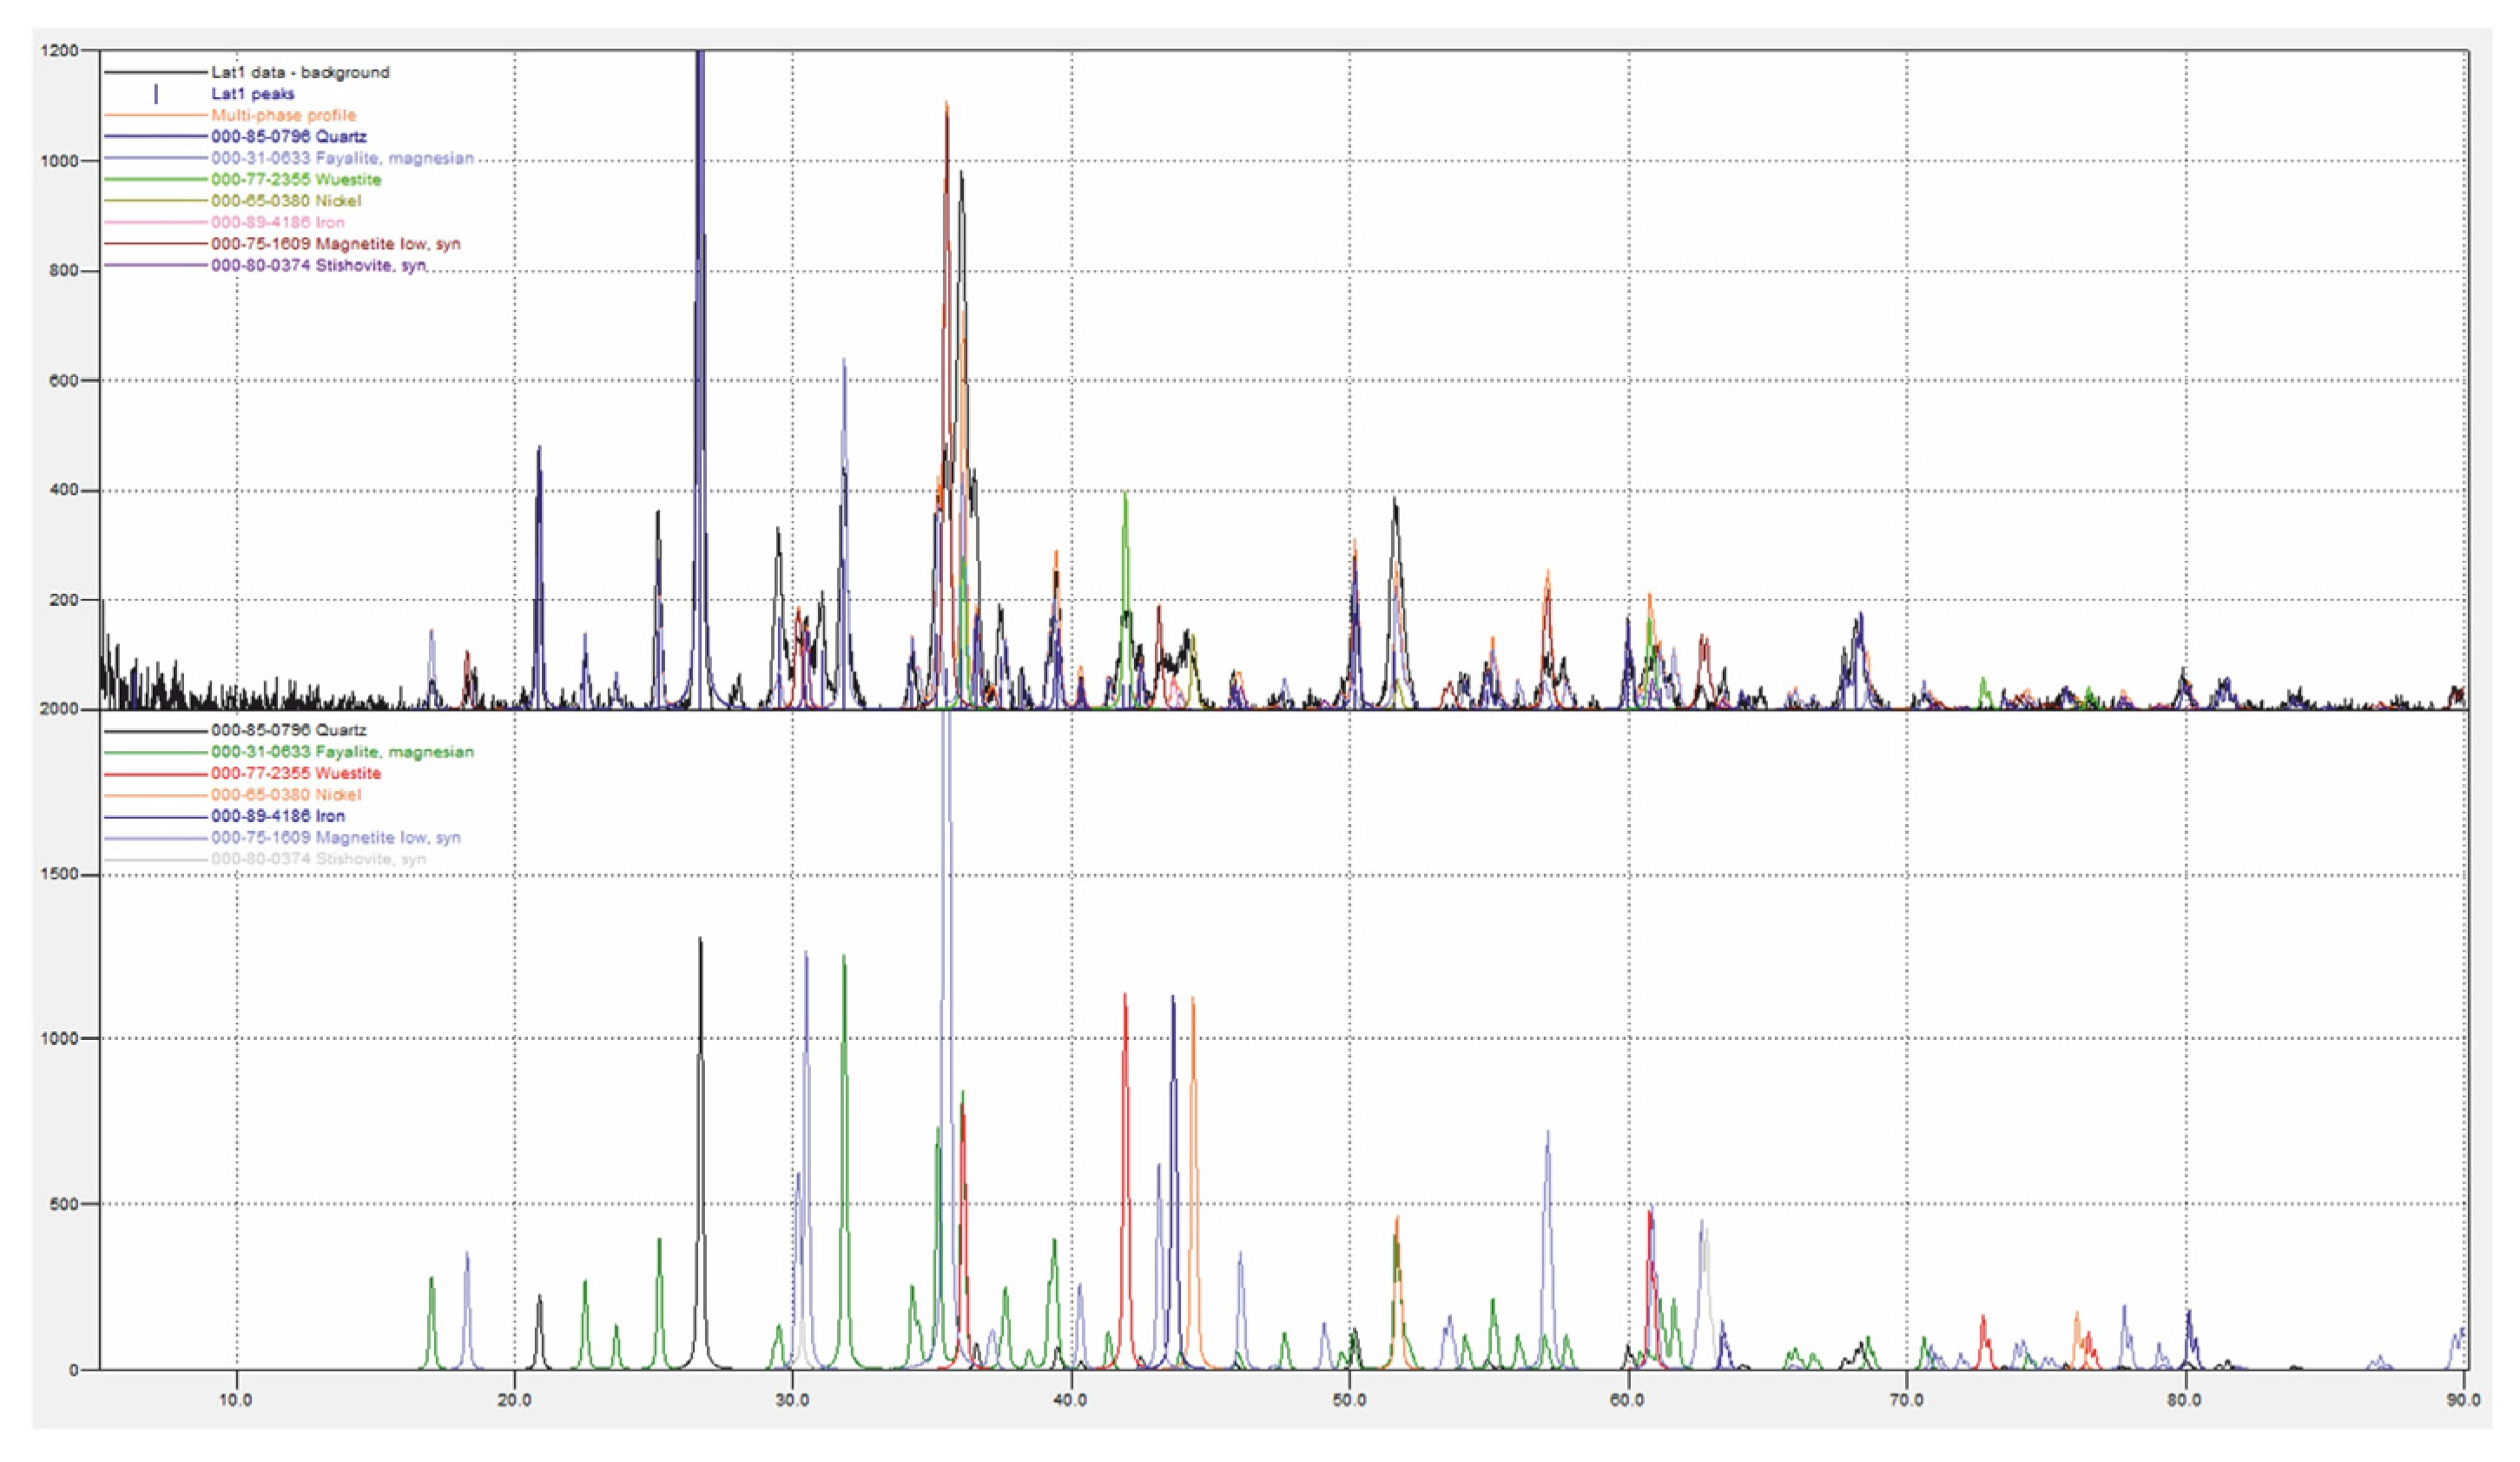
5 and ZnO were also identified. The measured loss on ignition (LOI) was only 5.39 wt.% indicating the presence of minor quantities of carbonates. XRD (
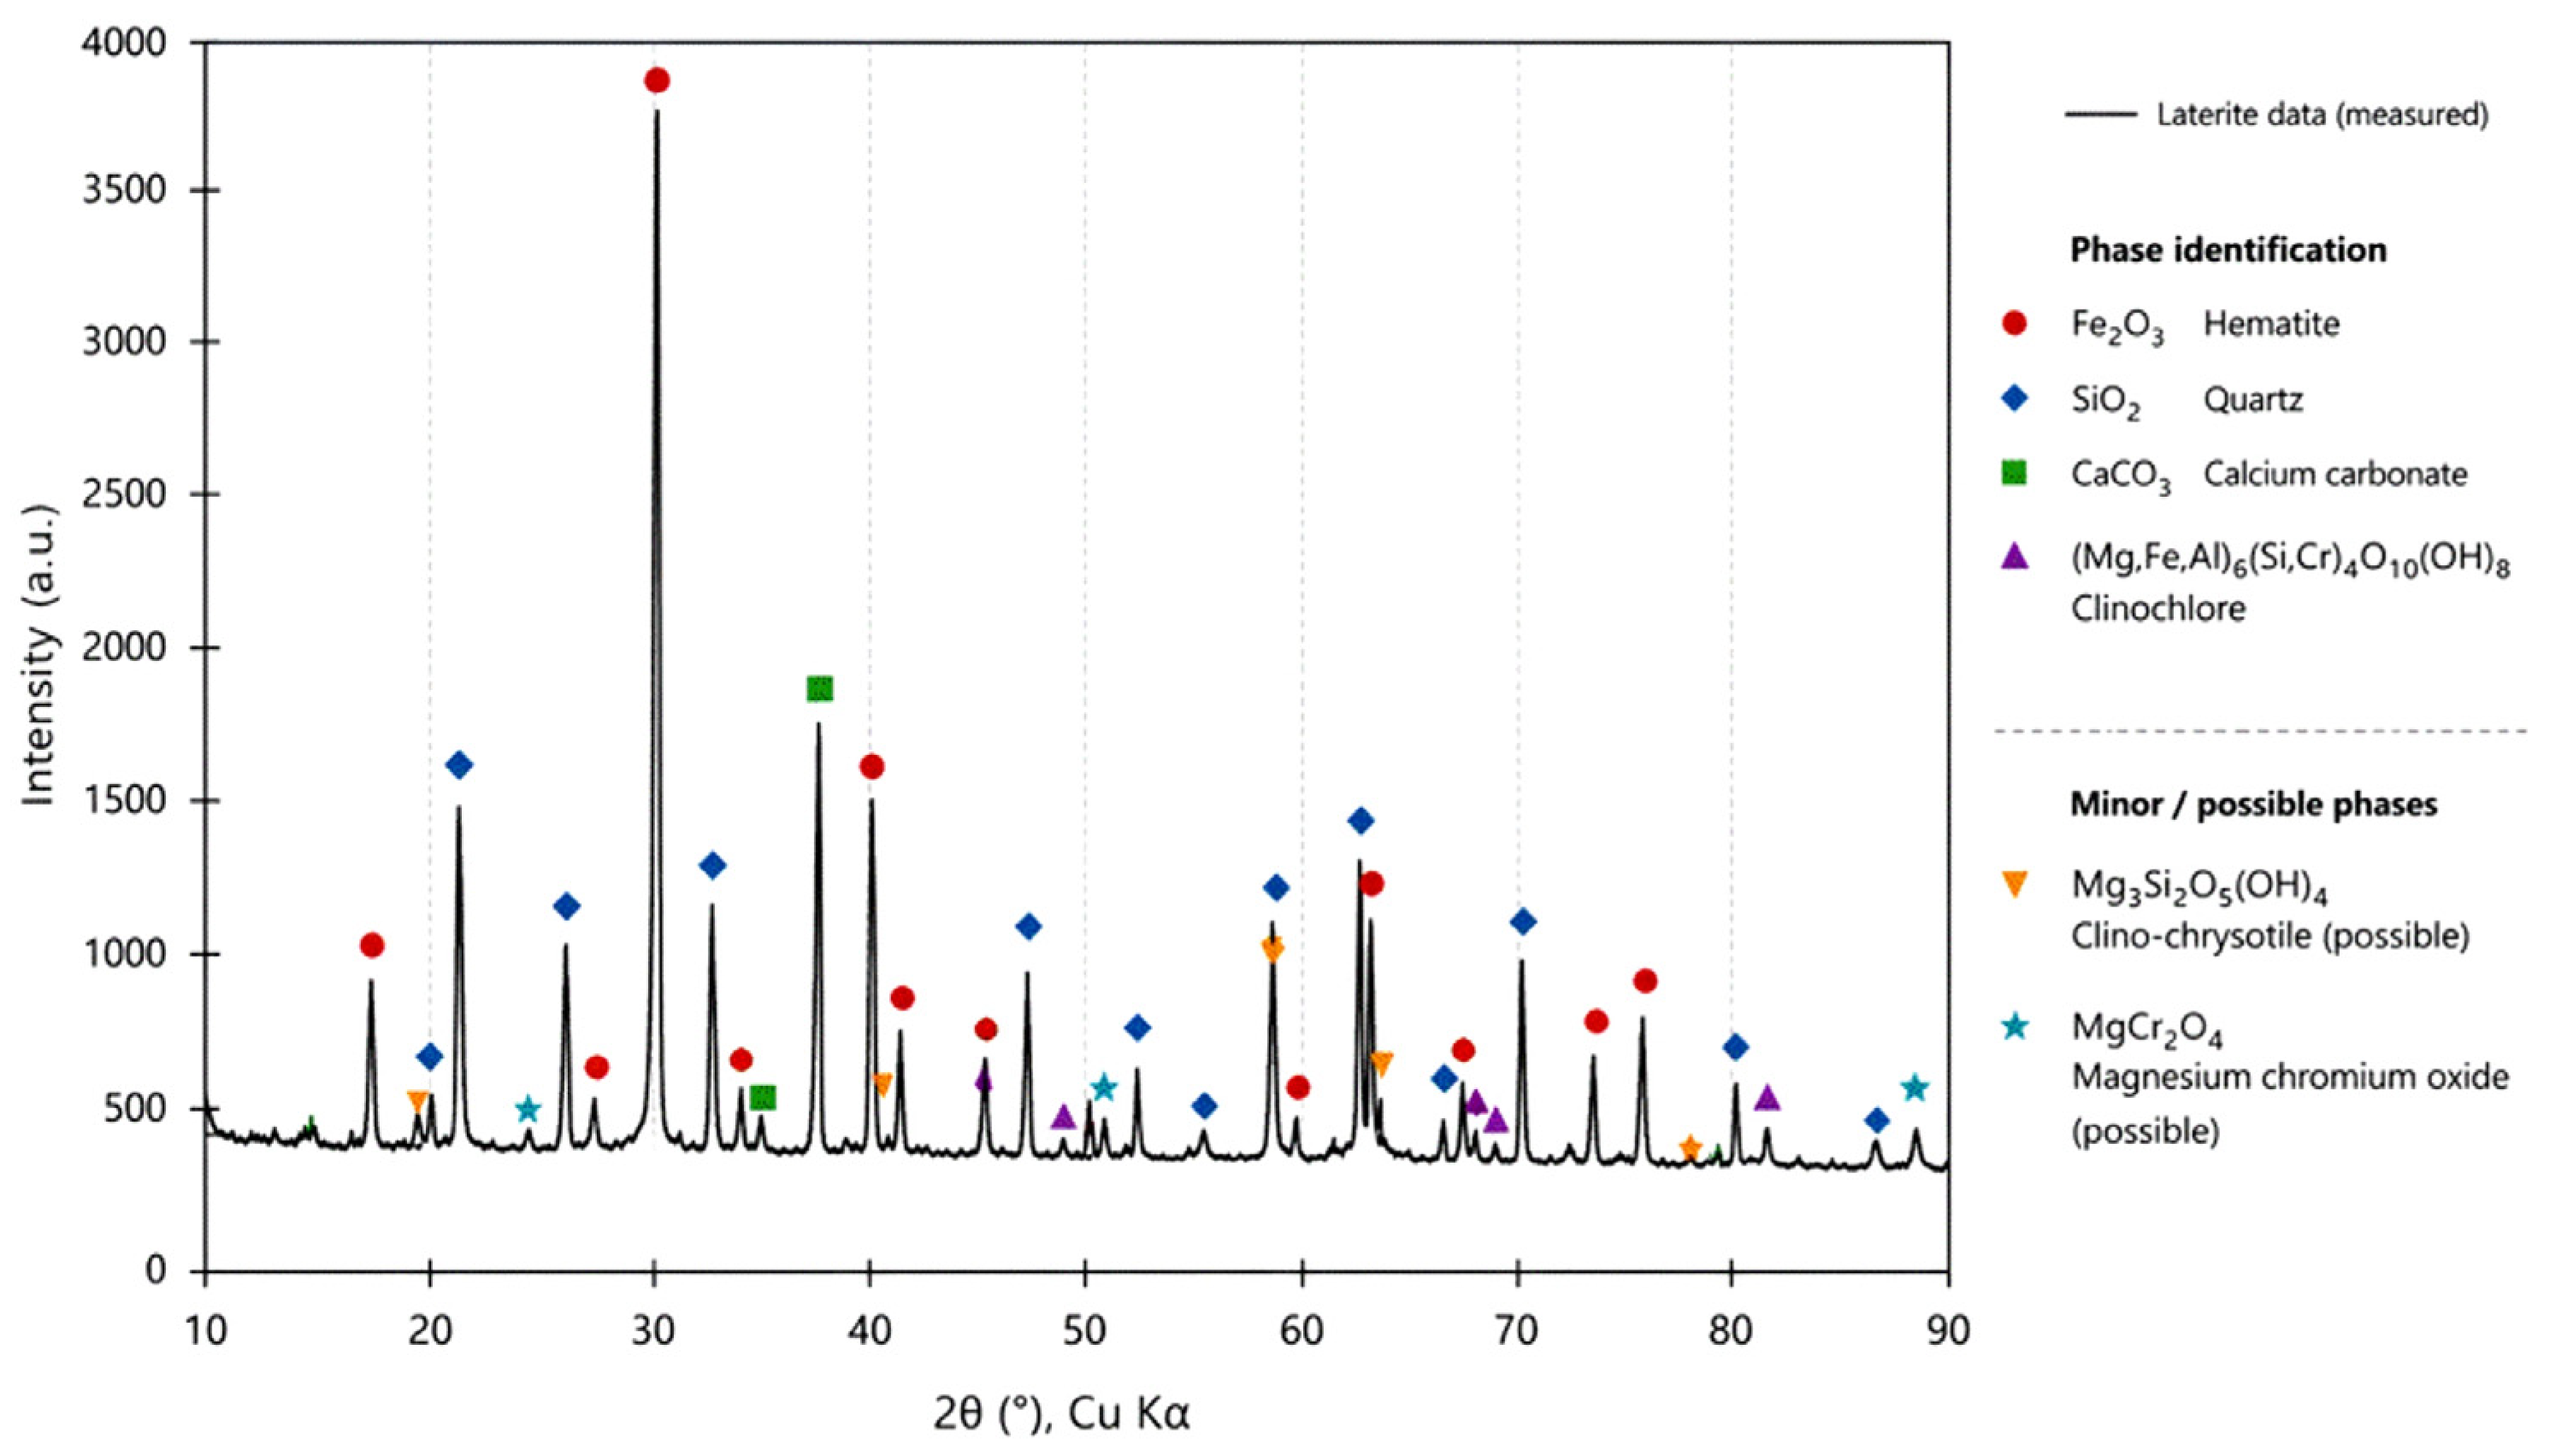
Figure 2) showed that the laterite is composed mainly of Fe
2O
3 (hematite), CaCO
3 (calcium carbonate), SiO
2 (quartz), (Mg,Fe,Al)
6(Si,Cr)
4O
10(OH)
8 (clinochlore). There was a possible presence of Mg
3Si
2O
5(OH)
4 (clinochrysotile) and MgCr
2O
4 (magnesium chromium oxide) as minor constituents. The particle-size distribution exhibited a relatively broad range, extending from below 0.25 mm to over 22.4 mm. Approximately 94% of the material passed the 22.4 mm sieve, while 73% was finer than 12.5 mm and 55% was below 7.1 mm. The proportion of fine particles gradually decreased towards smaller size fractions, with only 11.2% passing the 0.50 mm sieve and less than 5% passing the 0.25 mm sieve.
2.3. Laboratory IFRK and Instrumentation
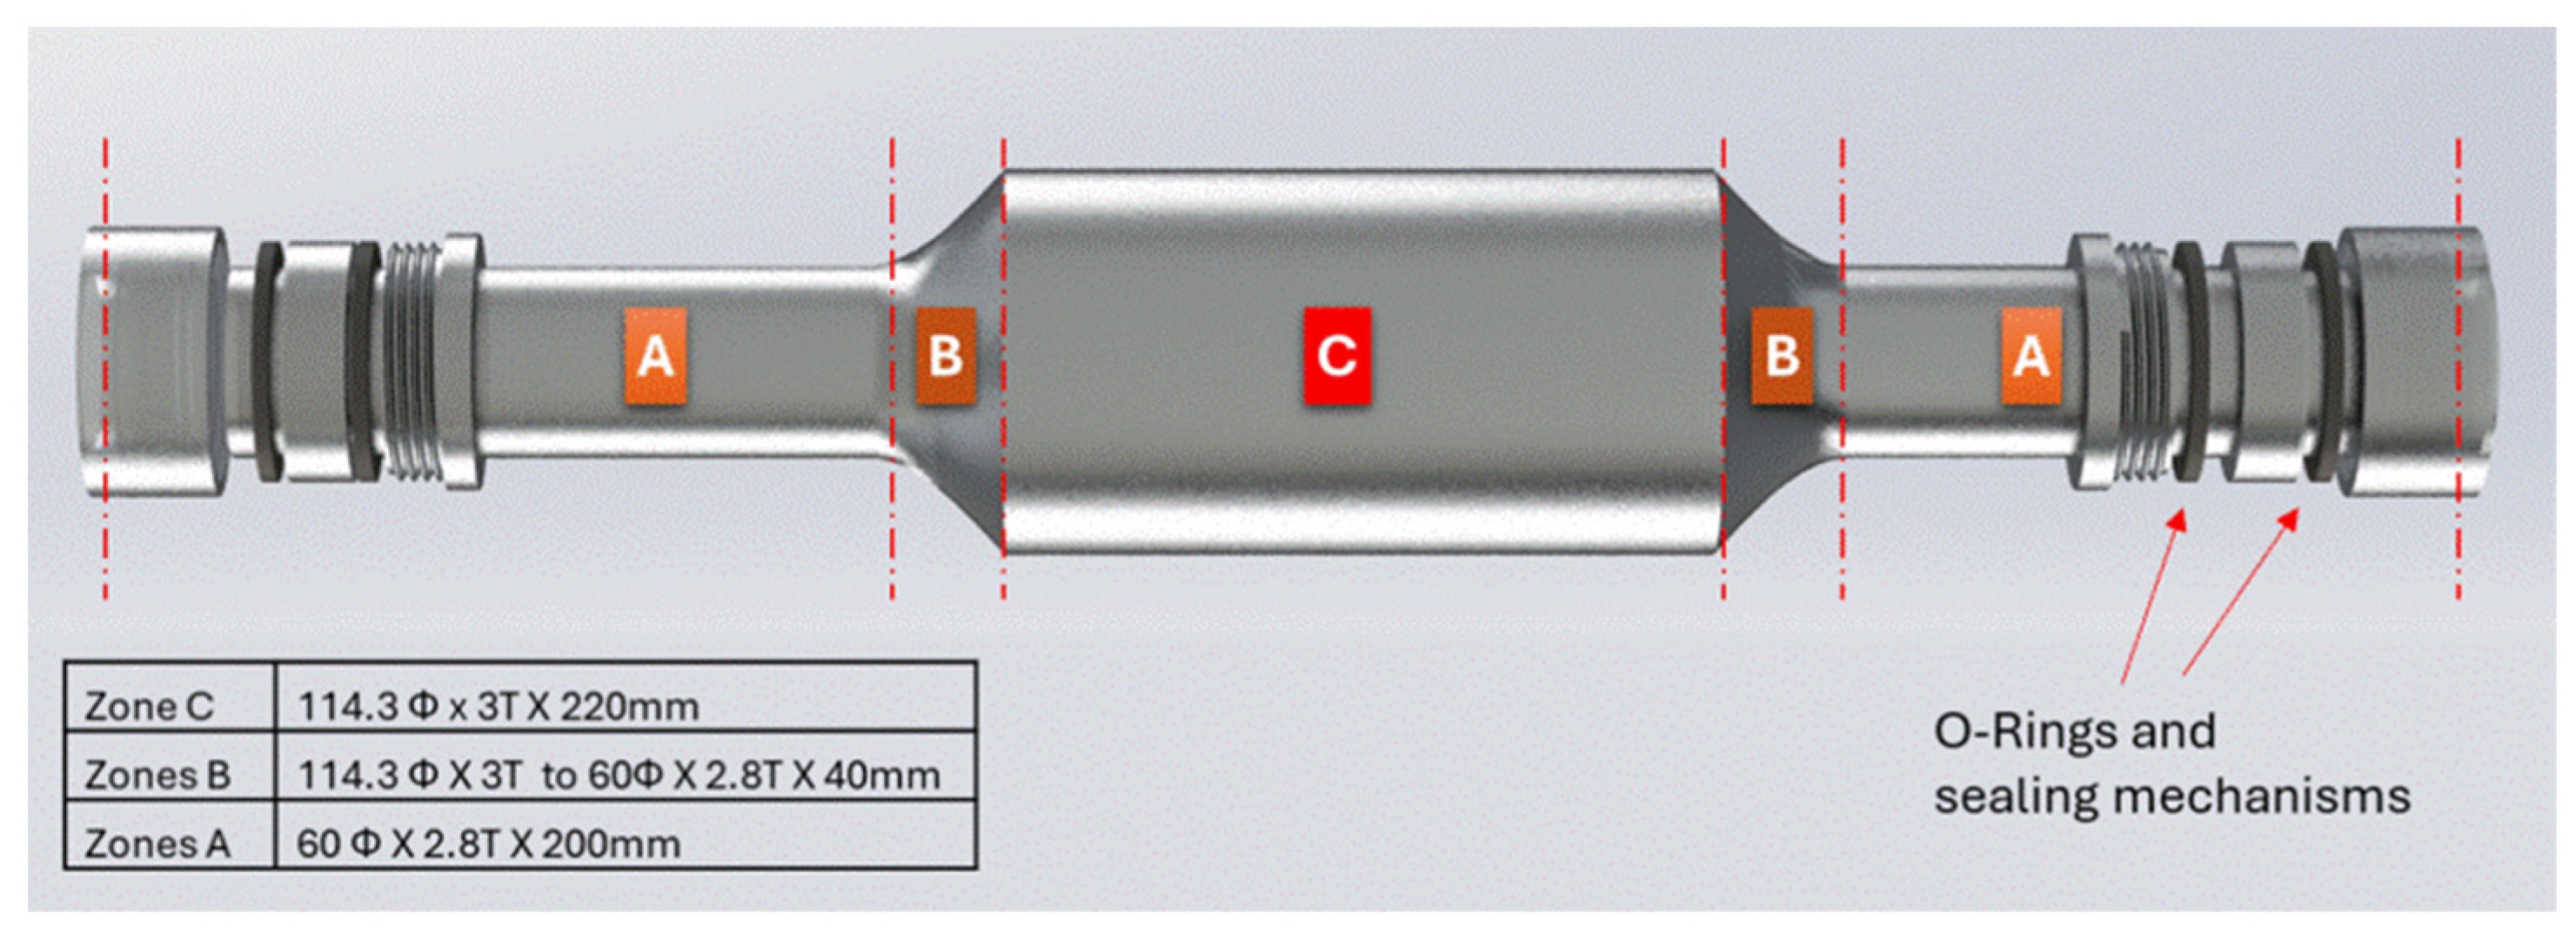
The experimental campaign was performed using a laboratory-scale electrically heated indirect-fired rotary kiln (IFRK) designed to provide precise control of both temperature and processing atmosphere during high-temperature treatment. The system was built around a horizontal rotary tube furnace, in which heat was transferred indirectly through electrically heated furnace walls rather than by direct contact between the heat source and the reacting material. This configuration ensured a uniform thermal environment while preventing contamination from combustion gases. The reactor tube was fabricated from heat-resistant STS310S stainless steel to withstand prolonged exposure to elevated temperatures and hydrogen-containing atmospheres. It uses a symmetrical three-section assembly to provide a well-defined reaction zone while ensuring stable gas flow and reliable sealing during operation (
Figure 3). The central section (Zone C) forms the main reaction chamber and is positioned entirely within the heated region of the furnace. On either side of the reactor, the central chamber is connected to the inlet and outlet tubes through two identical conical transition sections (Zones B). The inlet and outlet sections (Zones A) are straight cylindrical tubes which thermally separate the hot reaction chamber from the external connections, allowing instrumentation and gas fittings to operate at significantly lower temperatures. Inside the tube, four longitudinal lifters continuously mixed the material during rotation, improving particle movement and promoting more uniform heat distribution throughout the bed. Rotation was provided by a variable-speed drive operating between 1 and 10 rpm, while the furnace temperature was continuously monitored by a K-type thermocouple positioned in the hot zone. A PID controller maintained the desired thermal profile with high accuracy throughout each experimental run.
Figure 2.
XRD analysis in the lateritic ore sample.
Figure 2.
XRD analysis in the lateritic ore sample.
Figure 3.
IFRK cylindrical tube simulation in SOLIDWORKS 2026 [
31].
Figure 3.
IFRK cylindrical tube simulation in SOLIDWORKS 2026 [
31].
To preserve the controlled process atmosphere, both reactor ends were equipped with threaded sealing assemblies incorporating multiple high-temperature O-rings. This sealing arrangement was designed to accommodate thermal expansion while maintaining reactor tightness throughout rotation, effectively preventing air ingress and minimising process gas losses. The symmetric layout of the reactor also promotes balanced gas distribution and uniform thermal conditions, both of which are essential for reproducible calcination and hydrogen-reduction experiments.
2.4. Experimental Procedures
2.4.1. General Operating Procedure
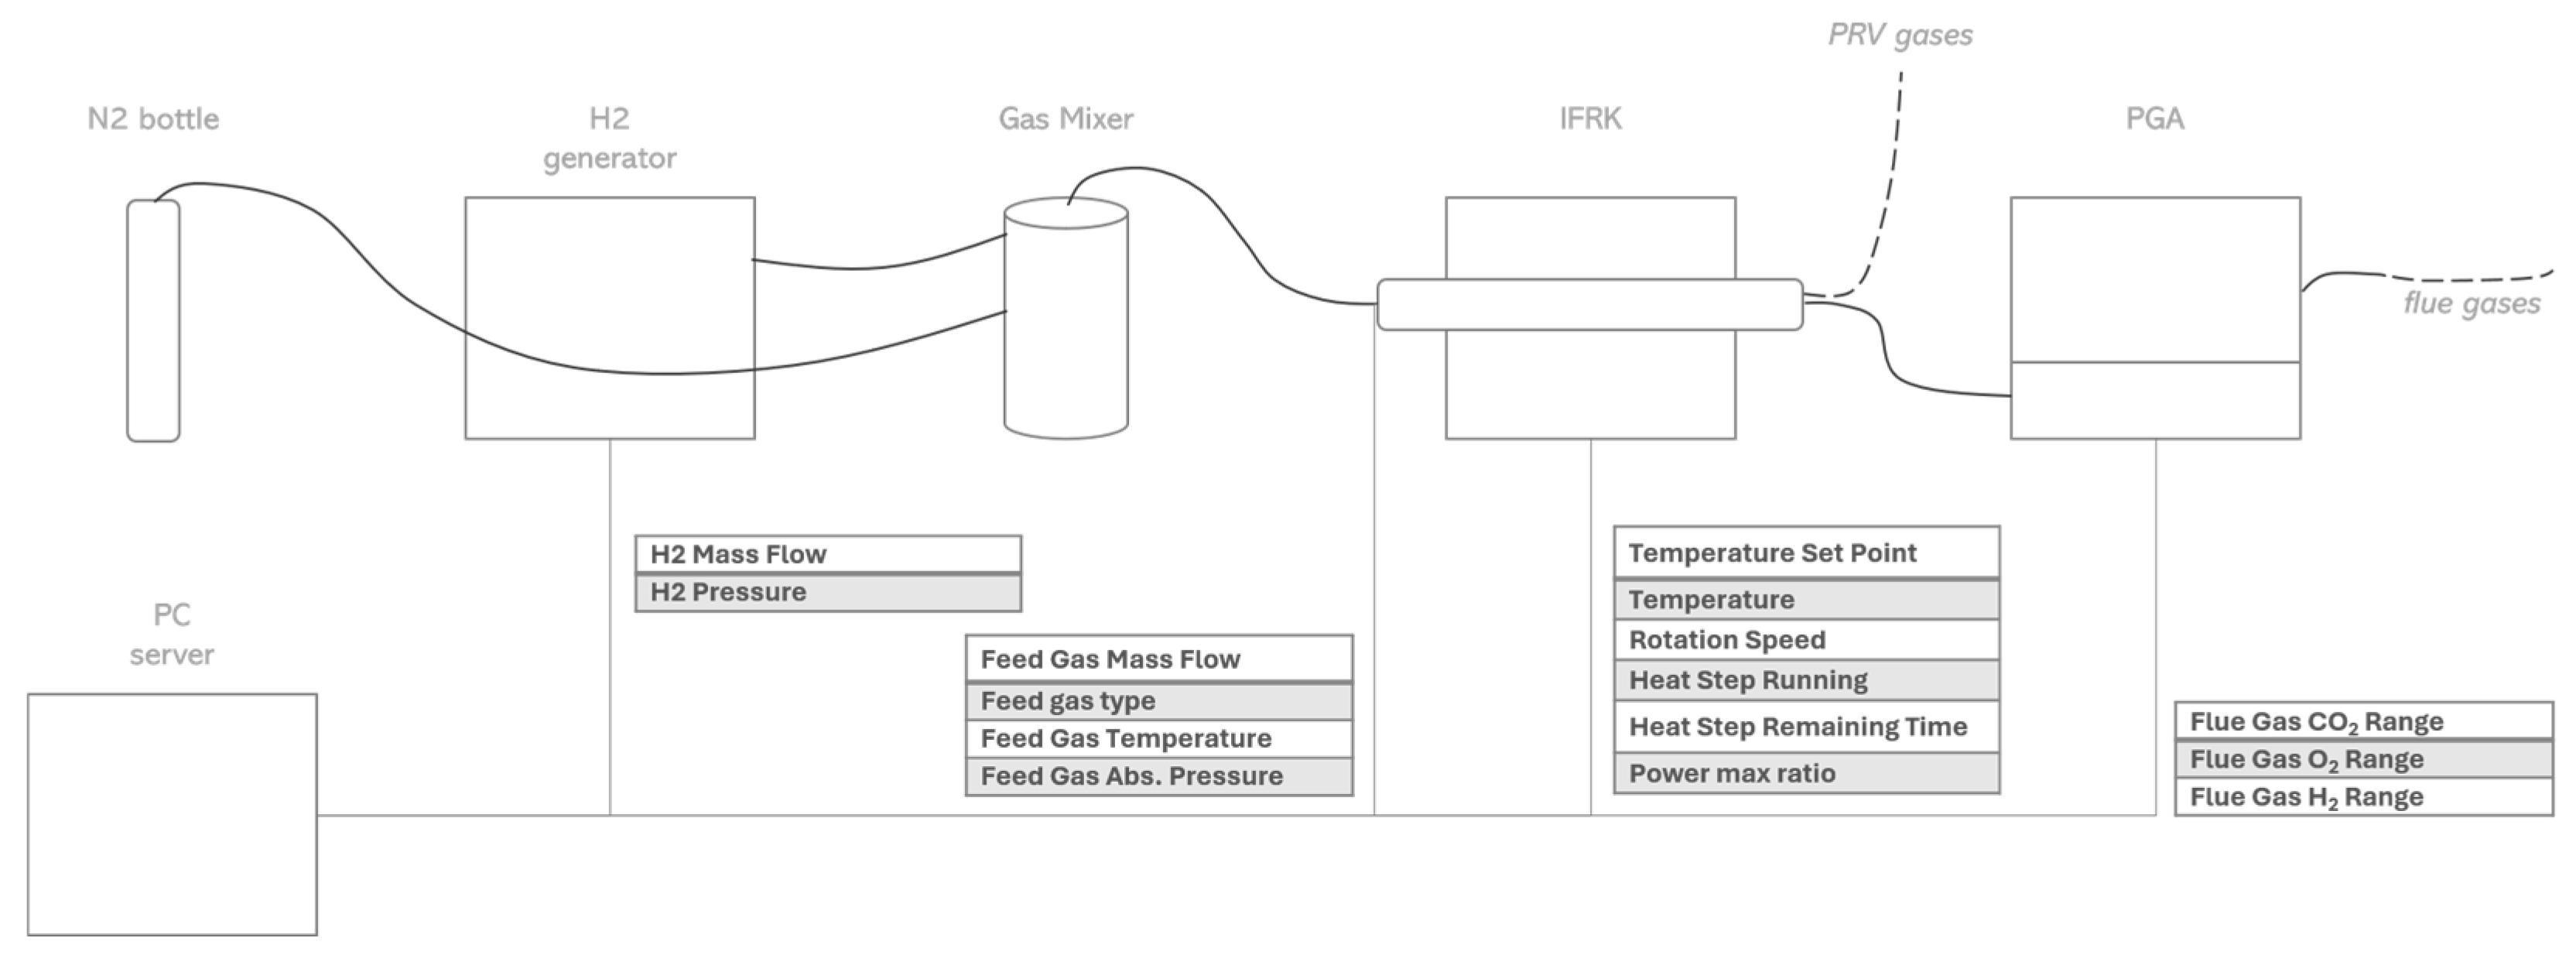
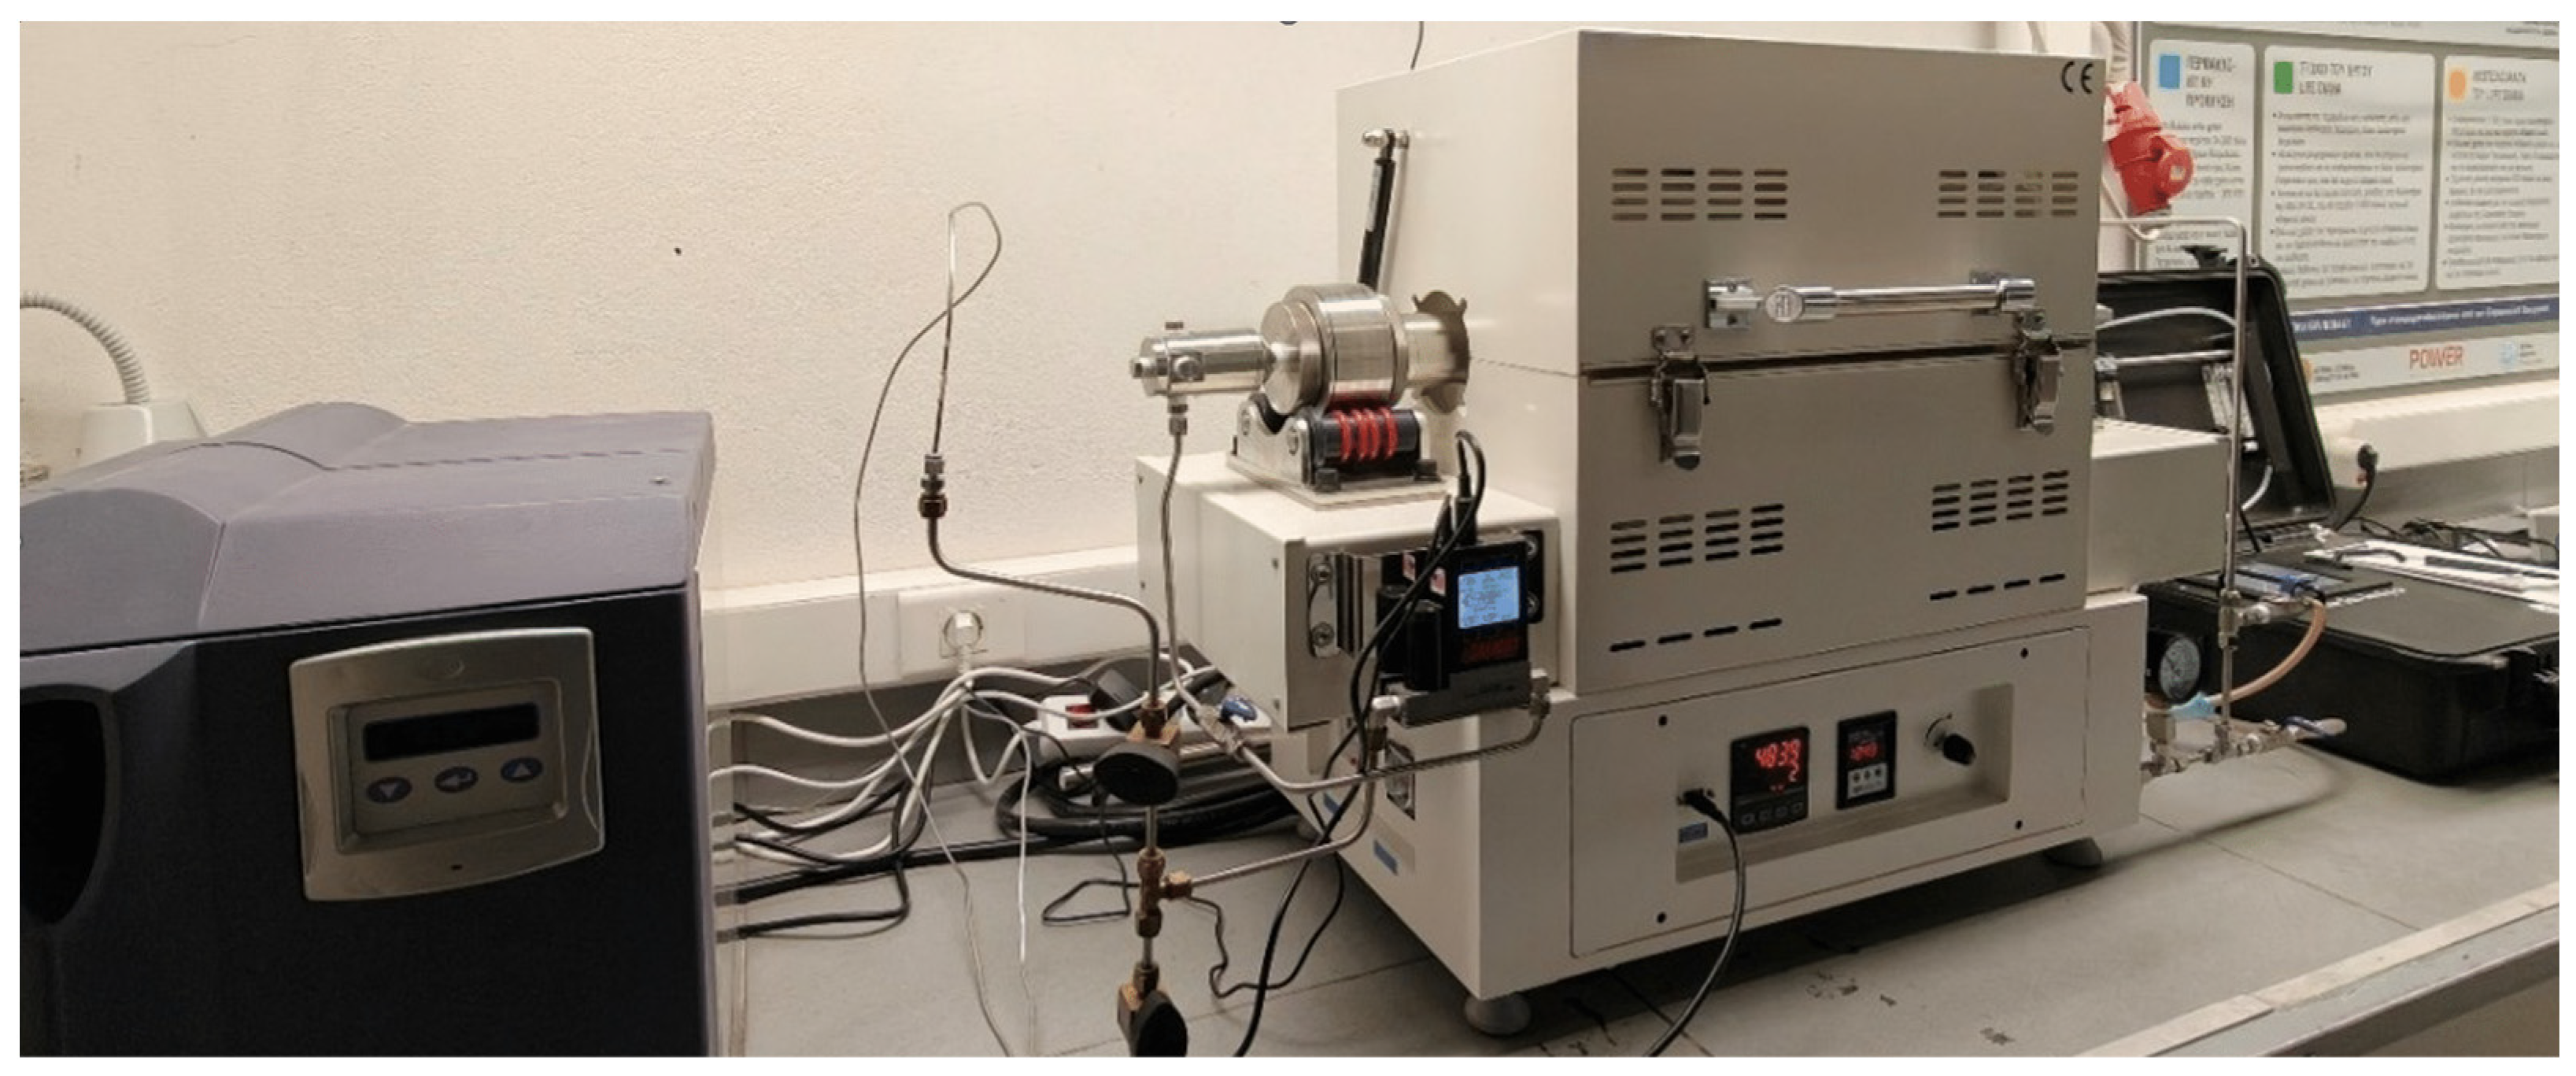
The experimental platform combined the IFRK with the gas supply network, atmosphere control system and data acquisition infrastructure, allowing all process variables to be monitored through a single integrated architecture (
Figure 4 and
Figure 5).
Nitrogen (99.999% purity) was supplied from a commercial compressed gas cylinder at an initial pressure of 200 bar and was used to purge the reactor before each experiment, establish an inert atmosphere during heating, and perform the final purge after the hydrogen-reduction tests. Hydrogen was generated on demand using a Parker 60H-MD proton exchange membrane (PEM) electrolyser, producing high-purity hydrogen (>99.999%). This approach eliminated the need for compressed hydrogen storage while ensuring a stable and continuous hydrogen supply throughout the reduction experiments. The hydrogen flow rate was regulated using a calibrated mass-flow controller before entering the sealed reactor. Once introduced into the sealed rotary reactor, the gas atmosphere was maintained under continuous control while the main operating parameters, including temperature, reactor rotation, gas-flow rate, inlet pressure and electrical power consumption, were recorded throughout each experimental run. The composition of the exhaust gas was monitored continuously using a portable infrared gas analyser (PGA-3510), enabling real-time measurement of gas composition.
The composition of the exhaust gas was continuously monitored throughout each experiment using a gas analyser connected directly to the reactor outlet. Since the reactor operated as a sealed system and the hydrogen inlet flow was accurately controlled by a calibrated mass-flow controller, the measured outlet hydrogen concentration was combined with the known inlet hydrogen flow to estimate hydrogen consumption during each reduction test. Although this approach was not intended to establish a complete mass balance, it provided a reliable and consistent basis for comparing hydrogen consumption under the different operating conditions examined in this work.
Particular attention was given to operational safety during commissioning. The laboratory was equipped with a dedicated mechanical ventilation system, fixed hydrogen leak detectors and portable gas detectors positioned around the installation to provide continuous leak monitoring. Before each experimental campaign, the reactor sealing system, gas distribution lines and all measurement devices were inspected, pressure-tested and calibrated to ensure stable and reproducible operation.
All process signals were collected through a central monitoring and data acquisition platform, enabling real-time visualisation and continuous recording of the experimental data. The commissioning procedure confirmed the integrity of the reactor, the stability of the gas handling system and the reliability of the monitoring equipment, providing a robust experimental platform for both magnesite calcination and hydrogen-assisted reduction under tightly controlled operating conditions.
During the experiments, the main operating variables—including furnace temperature and gas flow—were continuously regulated by the PID control system. Minor fluctuations around the selected setpoints were expected because of the thermal inertia of the furnace and the dynamic response of the gas supply system, and these were considered part of the normal operation of the experimental setup. Experimental measurements were recorded only after the operating conditions had stabilised, ensuring that the results presented in this work correspond to steady-state operation rather than transient start-up conditions.
Before the experimental programme commenced, the entire installation was commissioned to verify the performance of the reactor, gas handling system and monitoring equipment under both inert and hydrogen-based operating conditions. Nitrogen was supplied from a pressurised cylinder and used to purge the reactor before each experiment and to establish an inert atmosphere whenever required. Once introduced into the sealed rotary reactor, the gas atmosphere was maintained under continuous control while the main operating parameters, including temperature, reactor rotation, gas-flow rate, inlet pressure and electrical power consumption, were recorded throughout each experimental run.
2.4.2. Case Study A: Magnesite Calcination Experiments
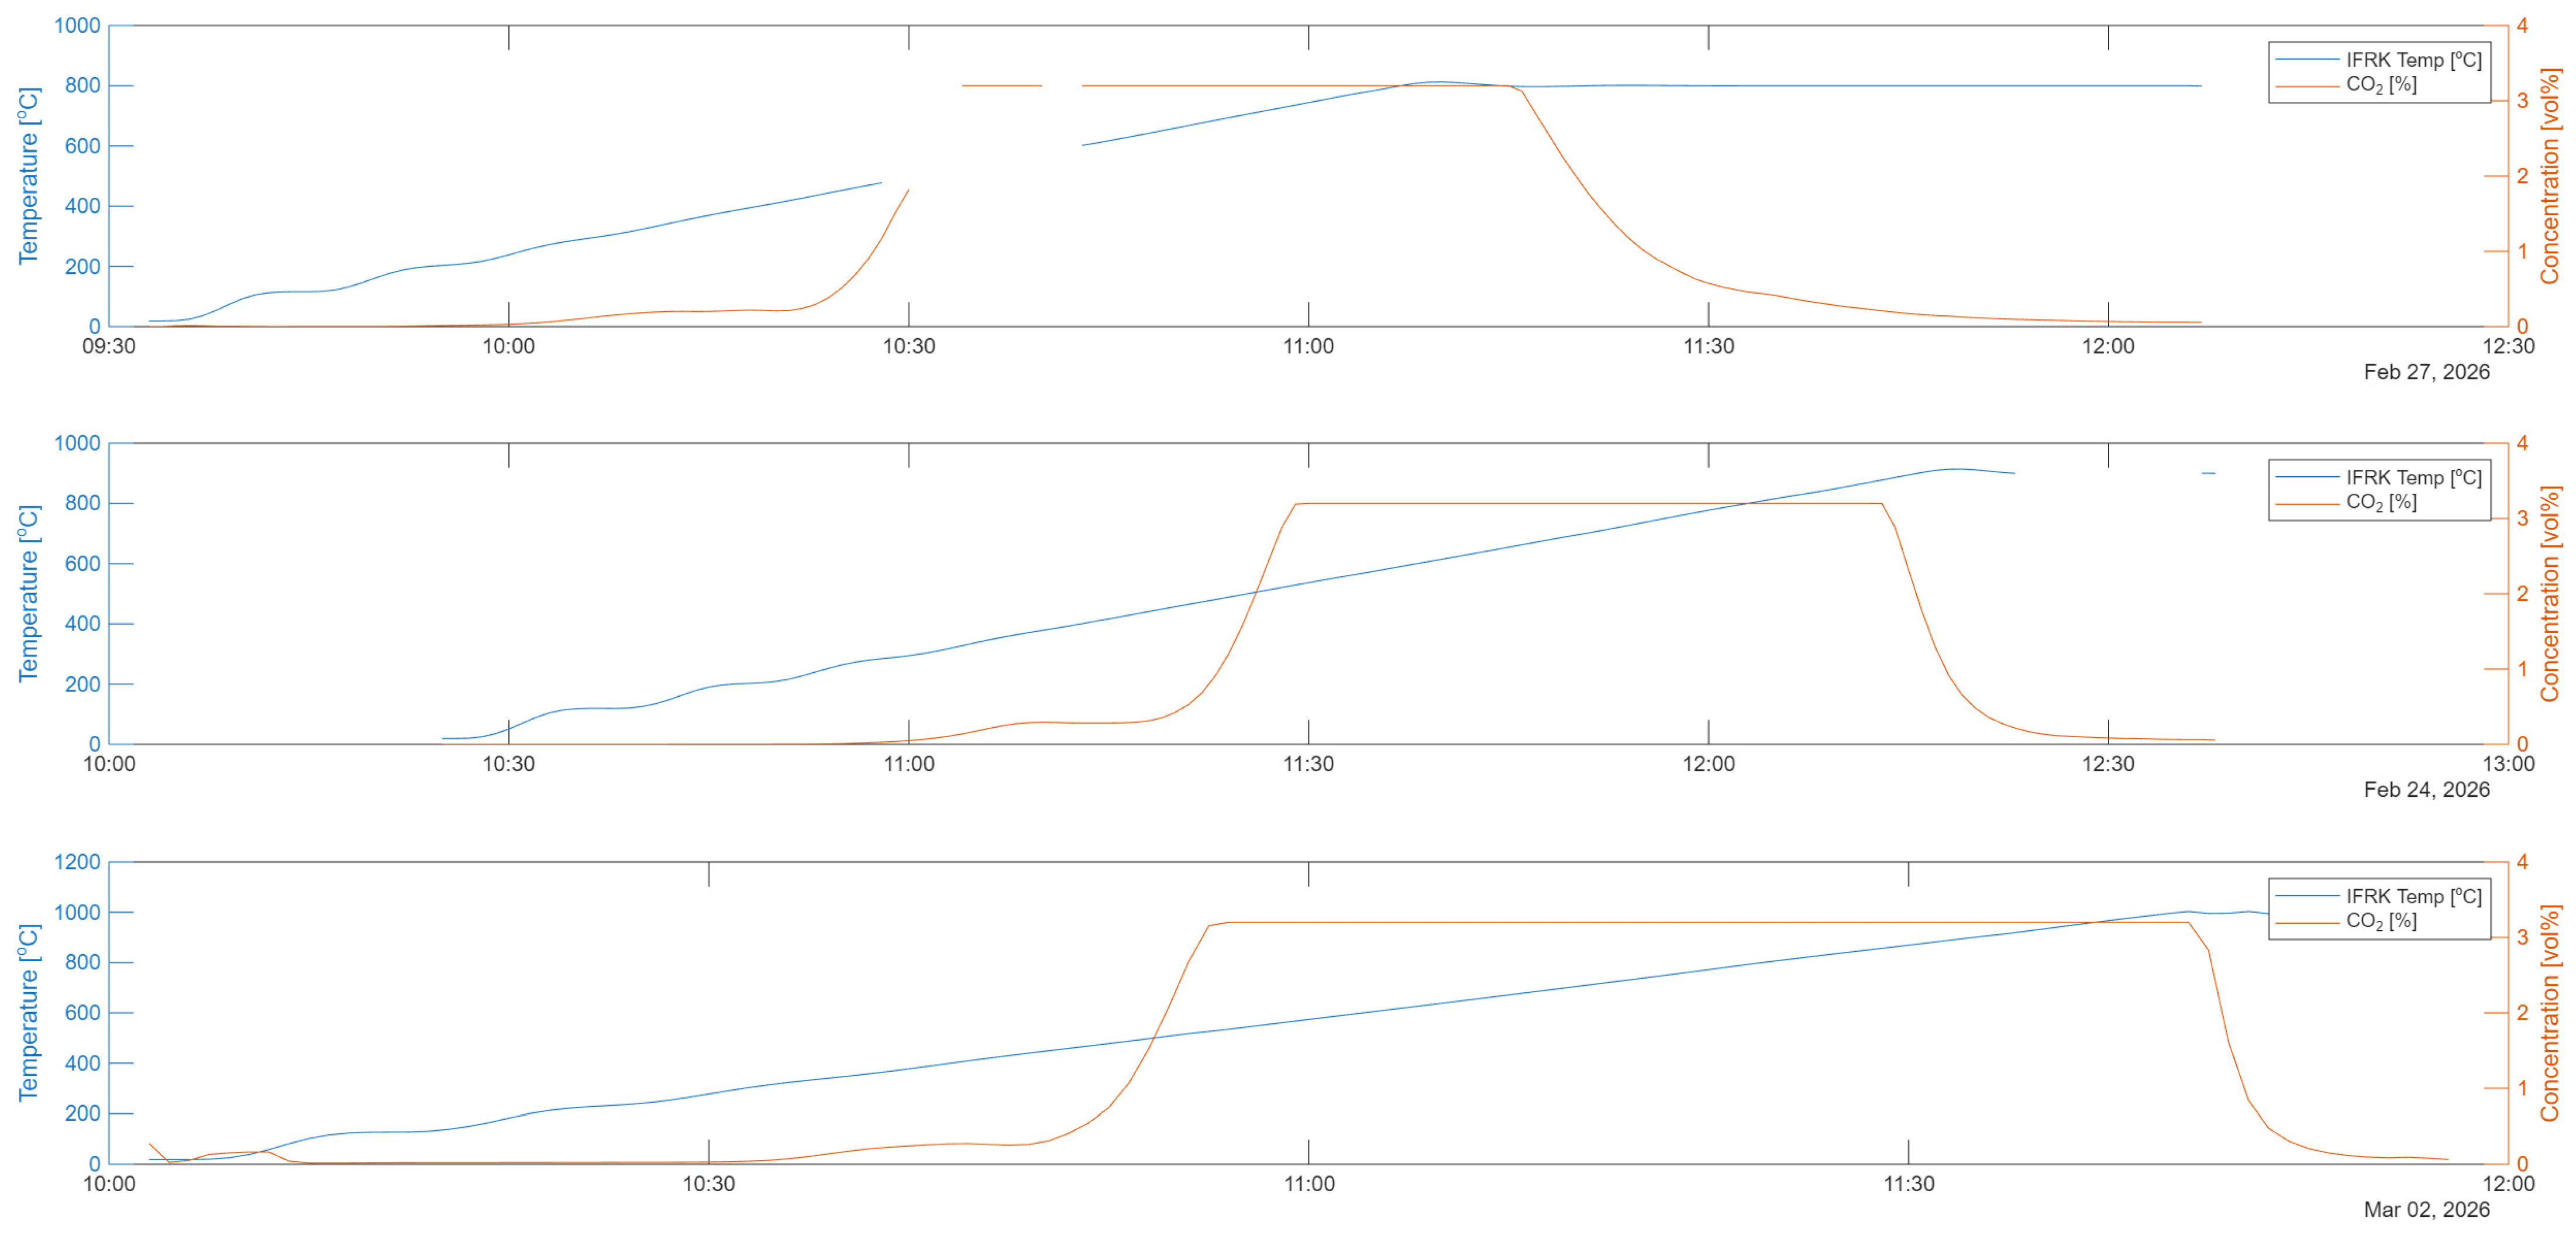
The experimental campaign was developed to investigate the calcination behaviour of magnesite in an airtight electrically heated indirect-fired rotary kiln over a range of representative operating temperatures. The study focused on the evolution of the decomposition process under controlled thermal conditions, with particular attention given to the relationship between temperature, reaction kinetics and residence time. The reactor was charged with the desired quantity of material, maintaining a filling degree below approximately 30% of the reaction chamber volume. This loading level ensured sufficient space for particle circulation and effective mixing during rotation. The reactor was operated at a constant rotational speed (10 rpm), allowing the internal lifters to continuously redistribute the material and maintain good contact between the particle bed and the heated reactor wall. Thermal treatment was performed at 800, 900 and 1000 °C, with temperature regulated automatically through the furnace PID controller at a nominal heating rate: approximately 8.0–9.8 °C min−1. A constant nitrogen flow was maintained during the entire thermal treatment to preserve the inert atmosphere and continuously remove the CO2 released during the decomposition of magnesite.
2.4.3. Case Study B: Laterite Hydrogen-Reduction Experiments
The experimental conditions were selected to evaluate the influence of the two parameters expected to have the greatest impact on hydrogen reduction, namely temperature and particle size, while keeping the remaining operating variables unchanged. Temperatures of 800 and 1000 °C were chosen to represent two characteristic operating regimes for hydrogen-assisted reduction, enabling the effect of increased thermal energy on the reduction process to be assessed. Two particle-size fractions were investigated to examine the influence of particle size on gas–solid interaction and reduction behaviour. Because reaction kinetics are significantly faster at higher temperatures, the hydrogen-treatment time was set to 35 min at 800 °C and reduced to 15 min at 1000 °C to achieve representative reduction conditions without unnecessarily extending the experiments. Throughout the study, the reactor rotation speed, hydrogen flow rate and all other operating conditions were maintained constant so that the observed differences could be attributed primarily to the selected experimental variables. During all tests, the reactor rotated at 10 rpm, providing continuous movement of the particle bed and promoting uniform contact between the material and the heated reactor surface.
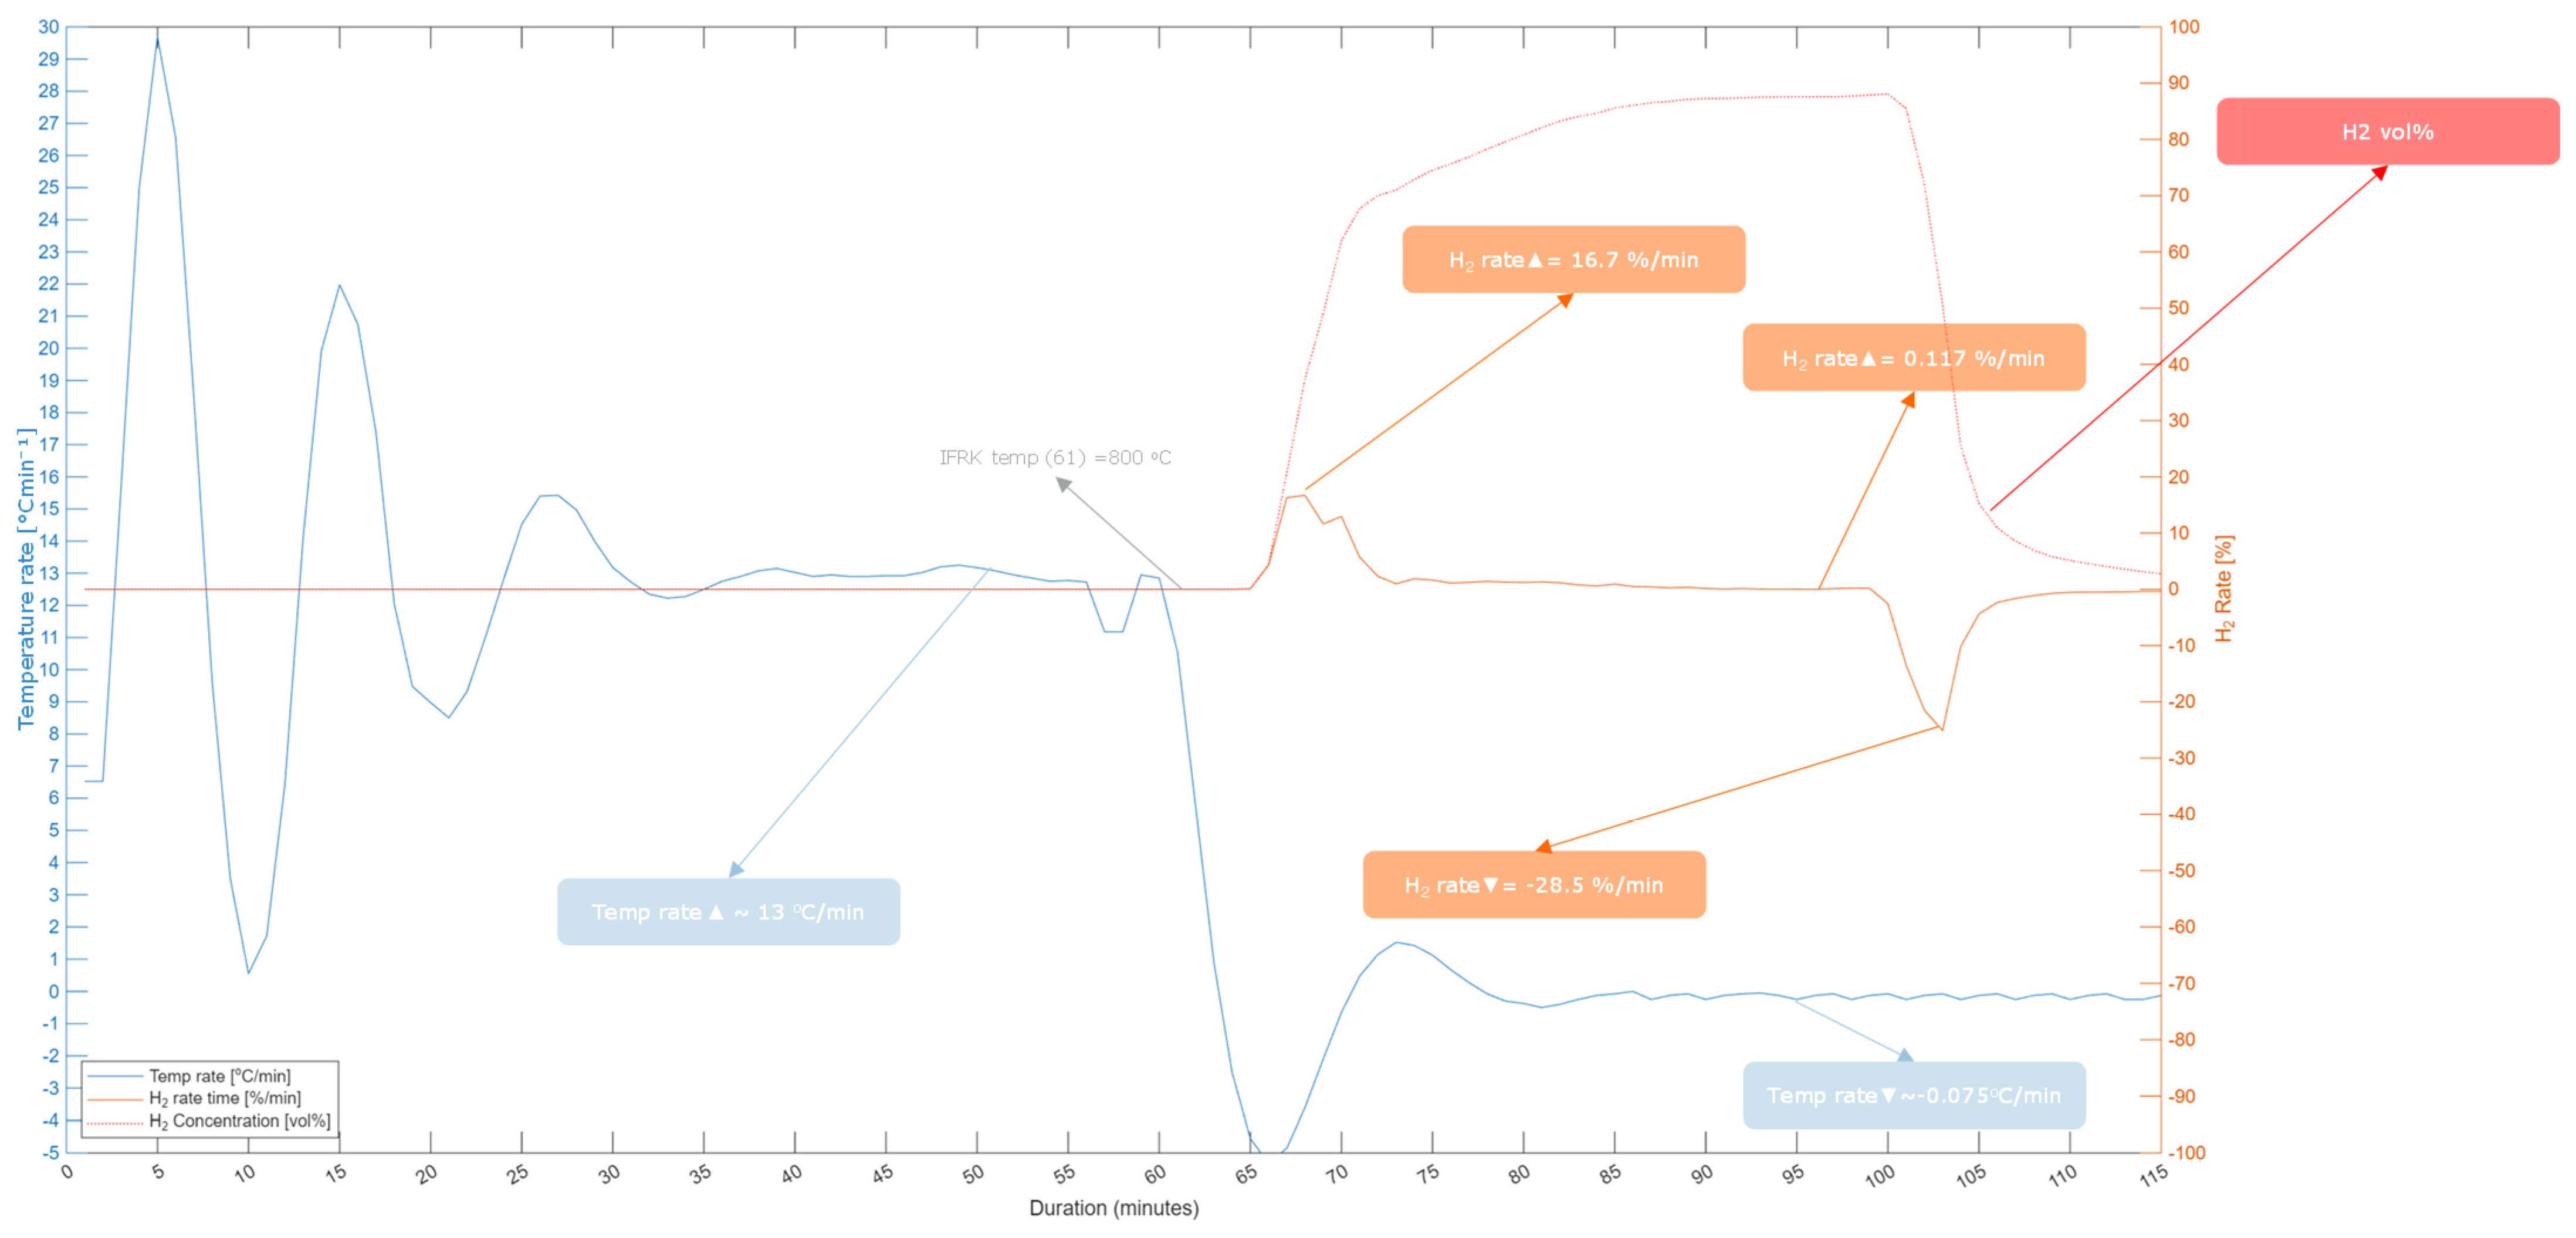
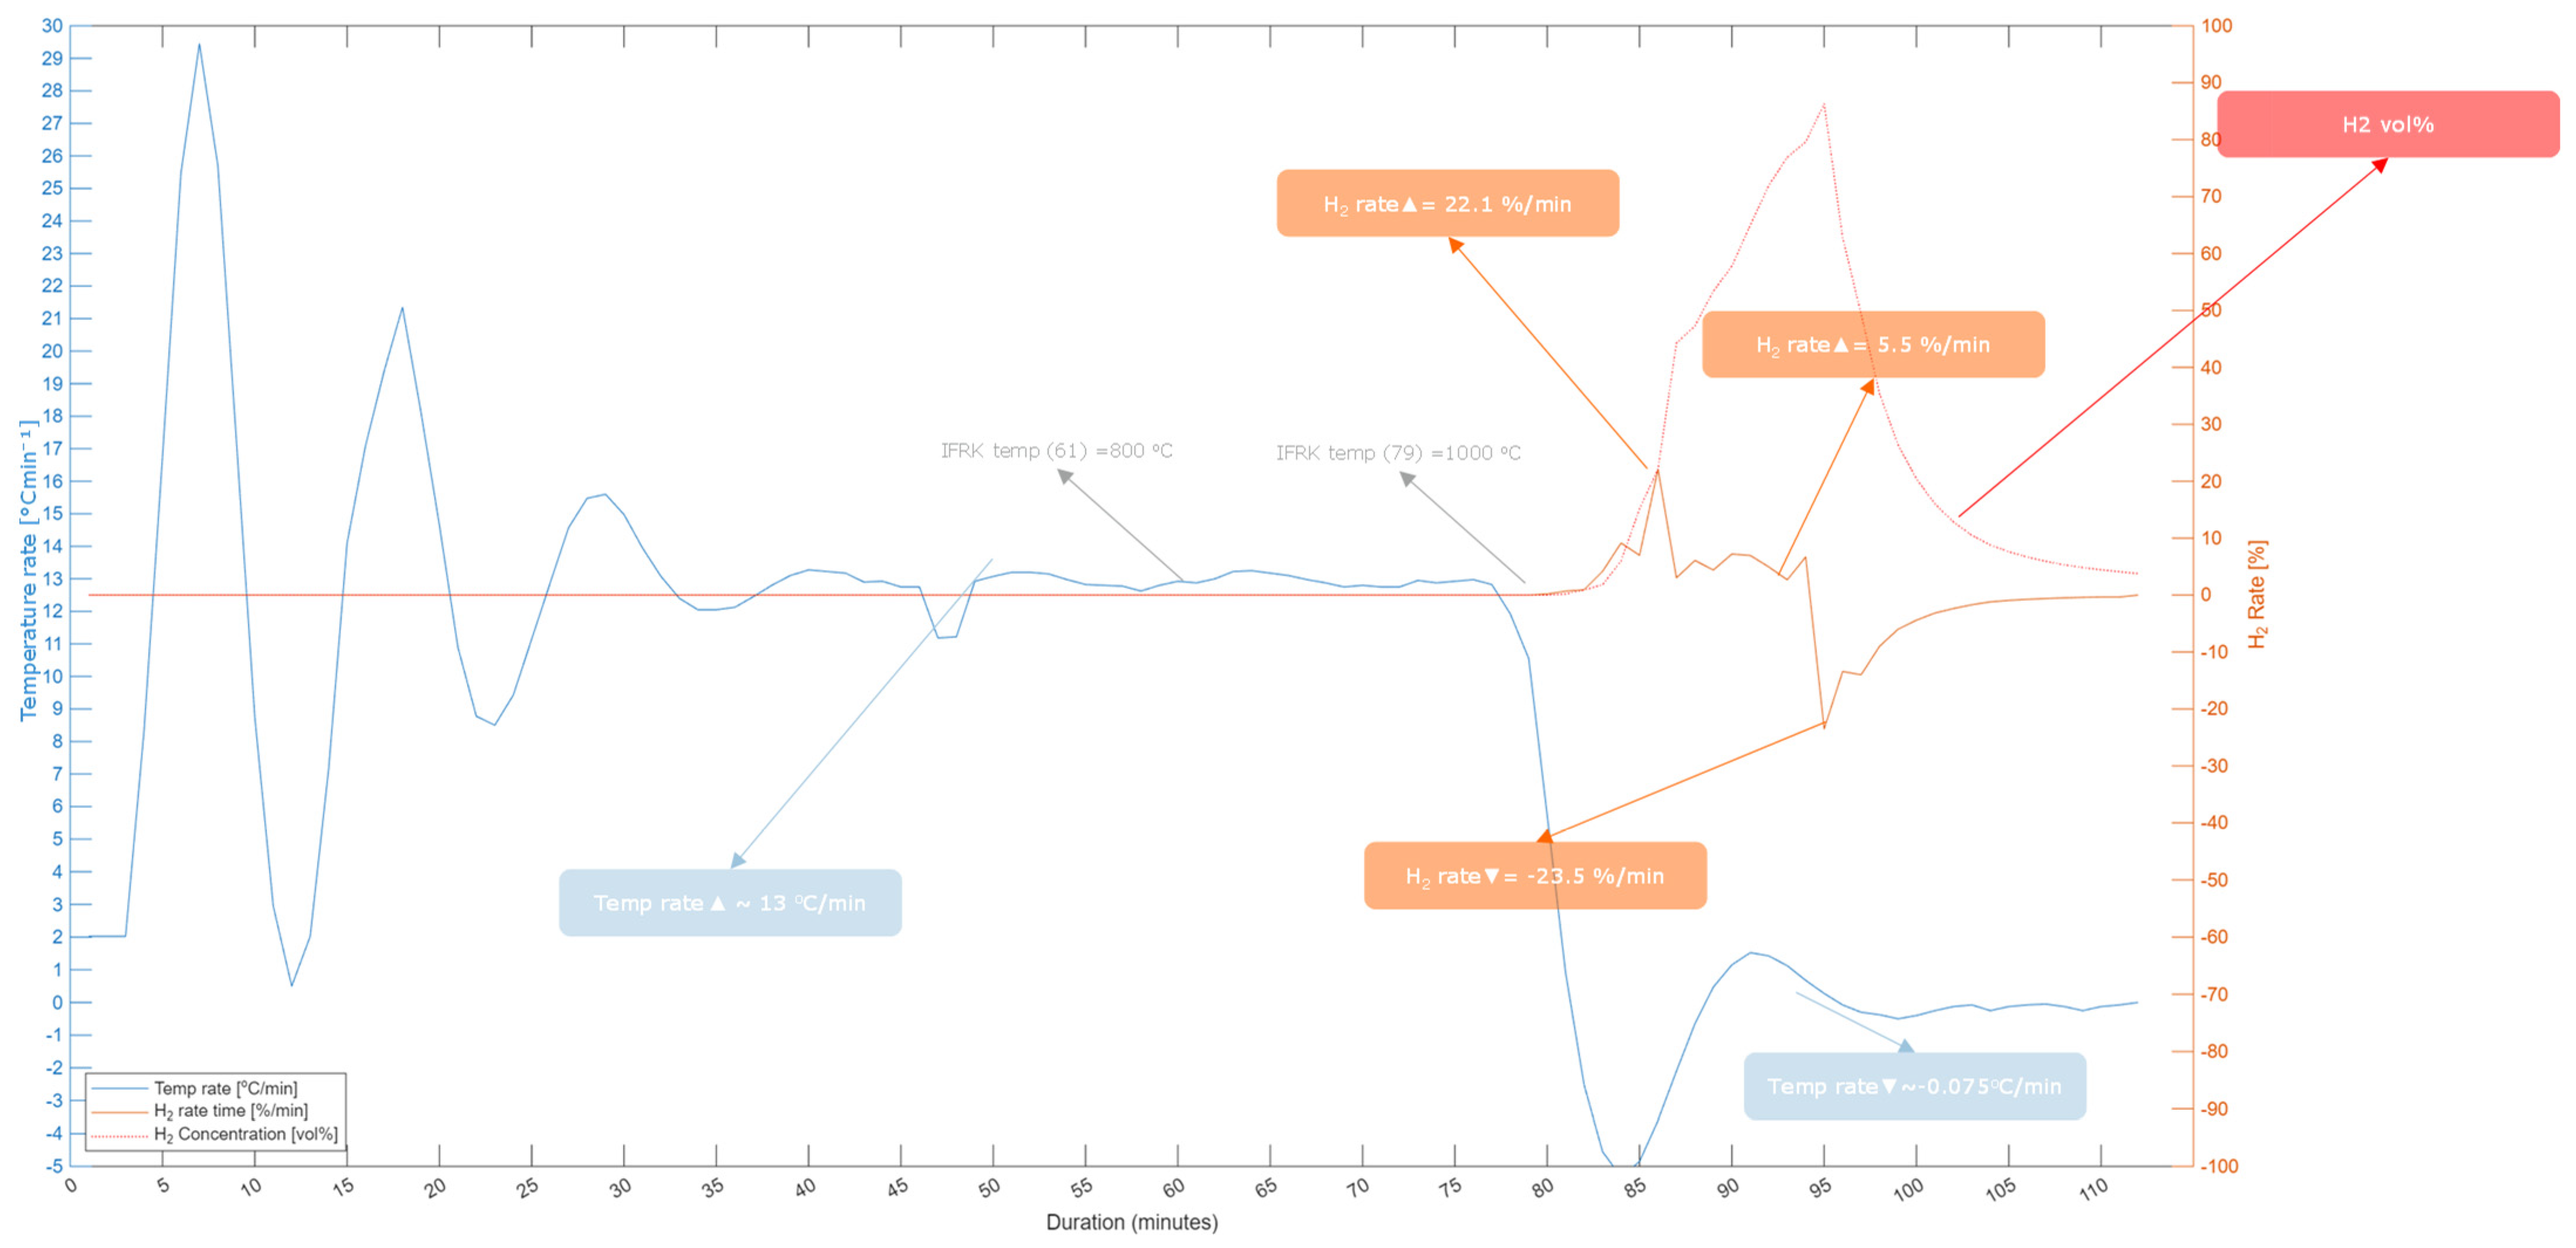
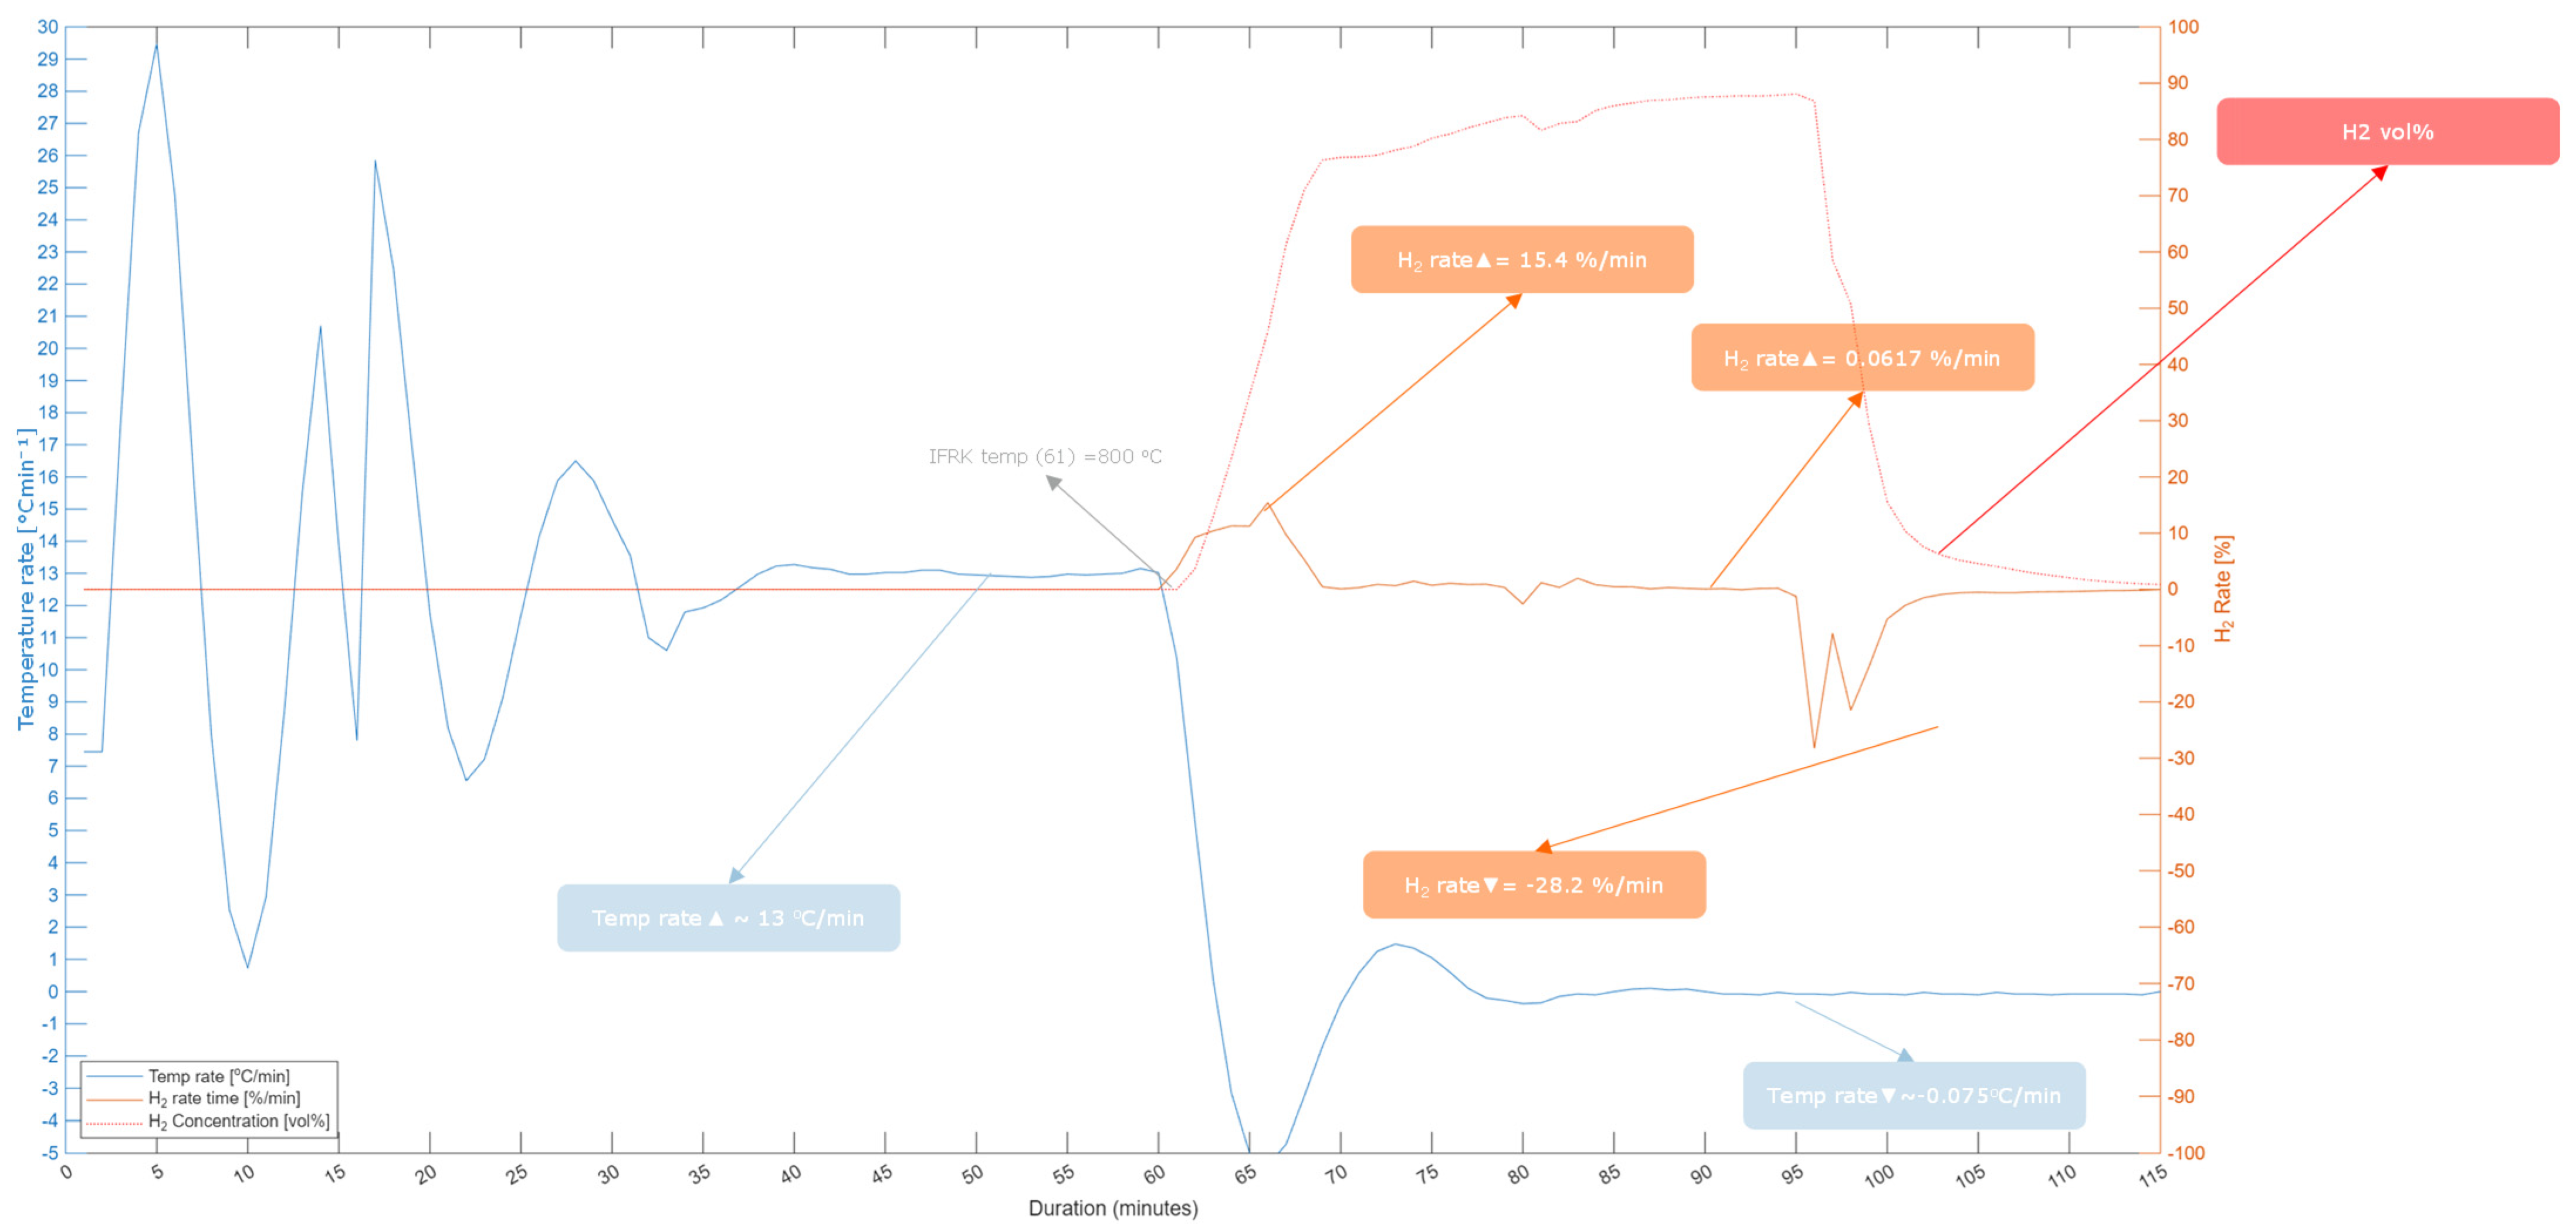
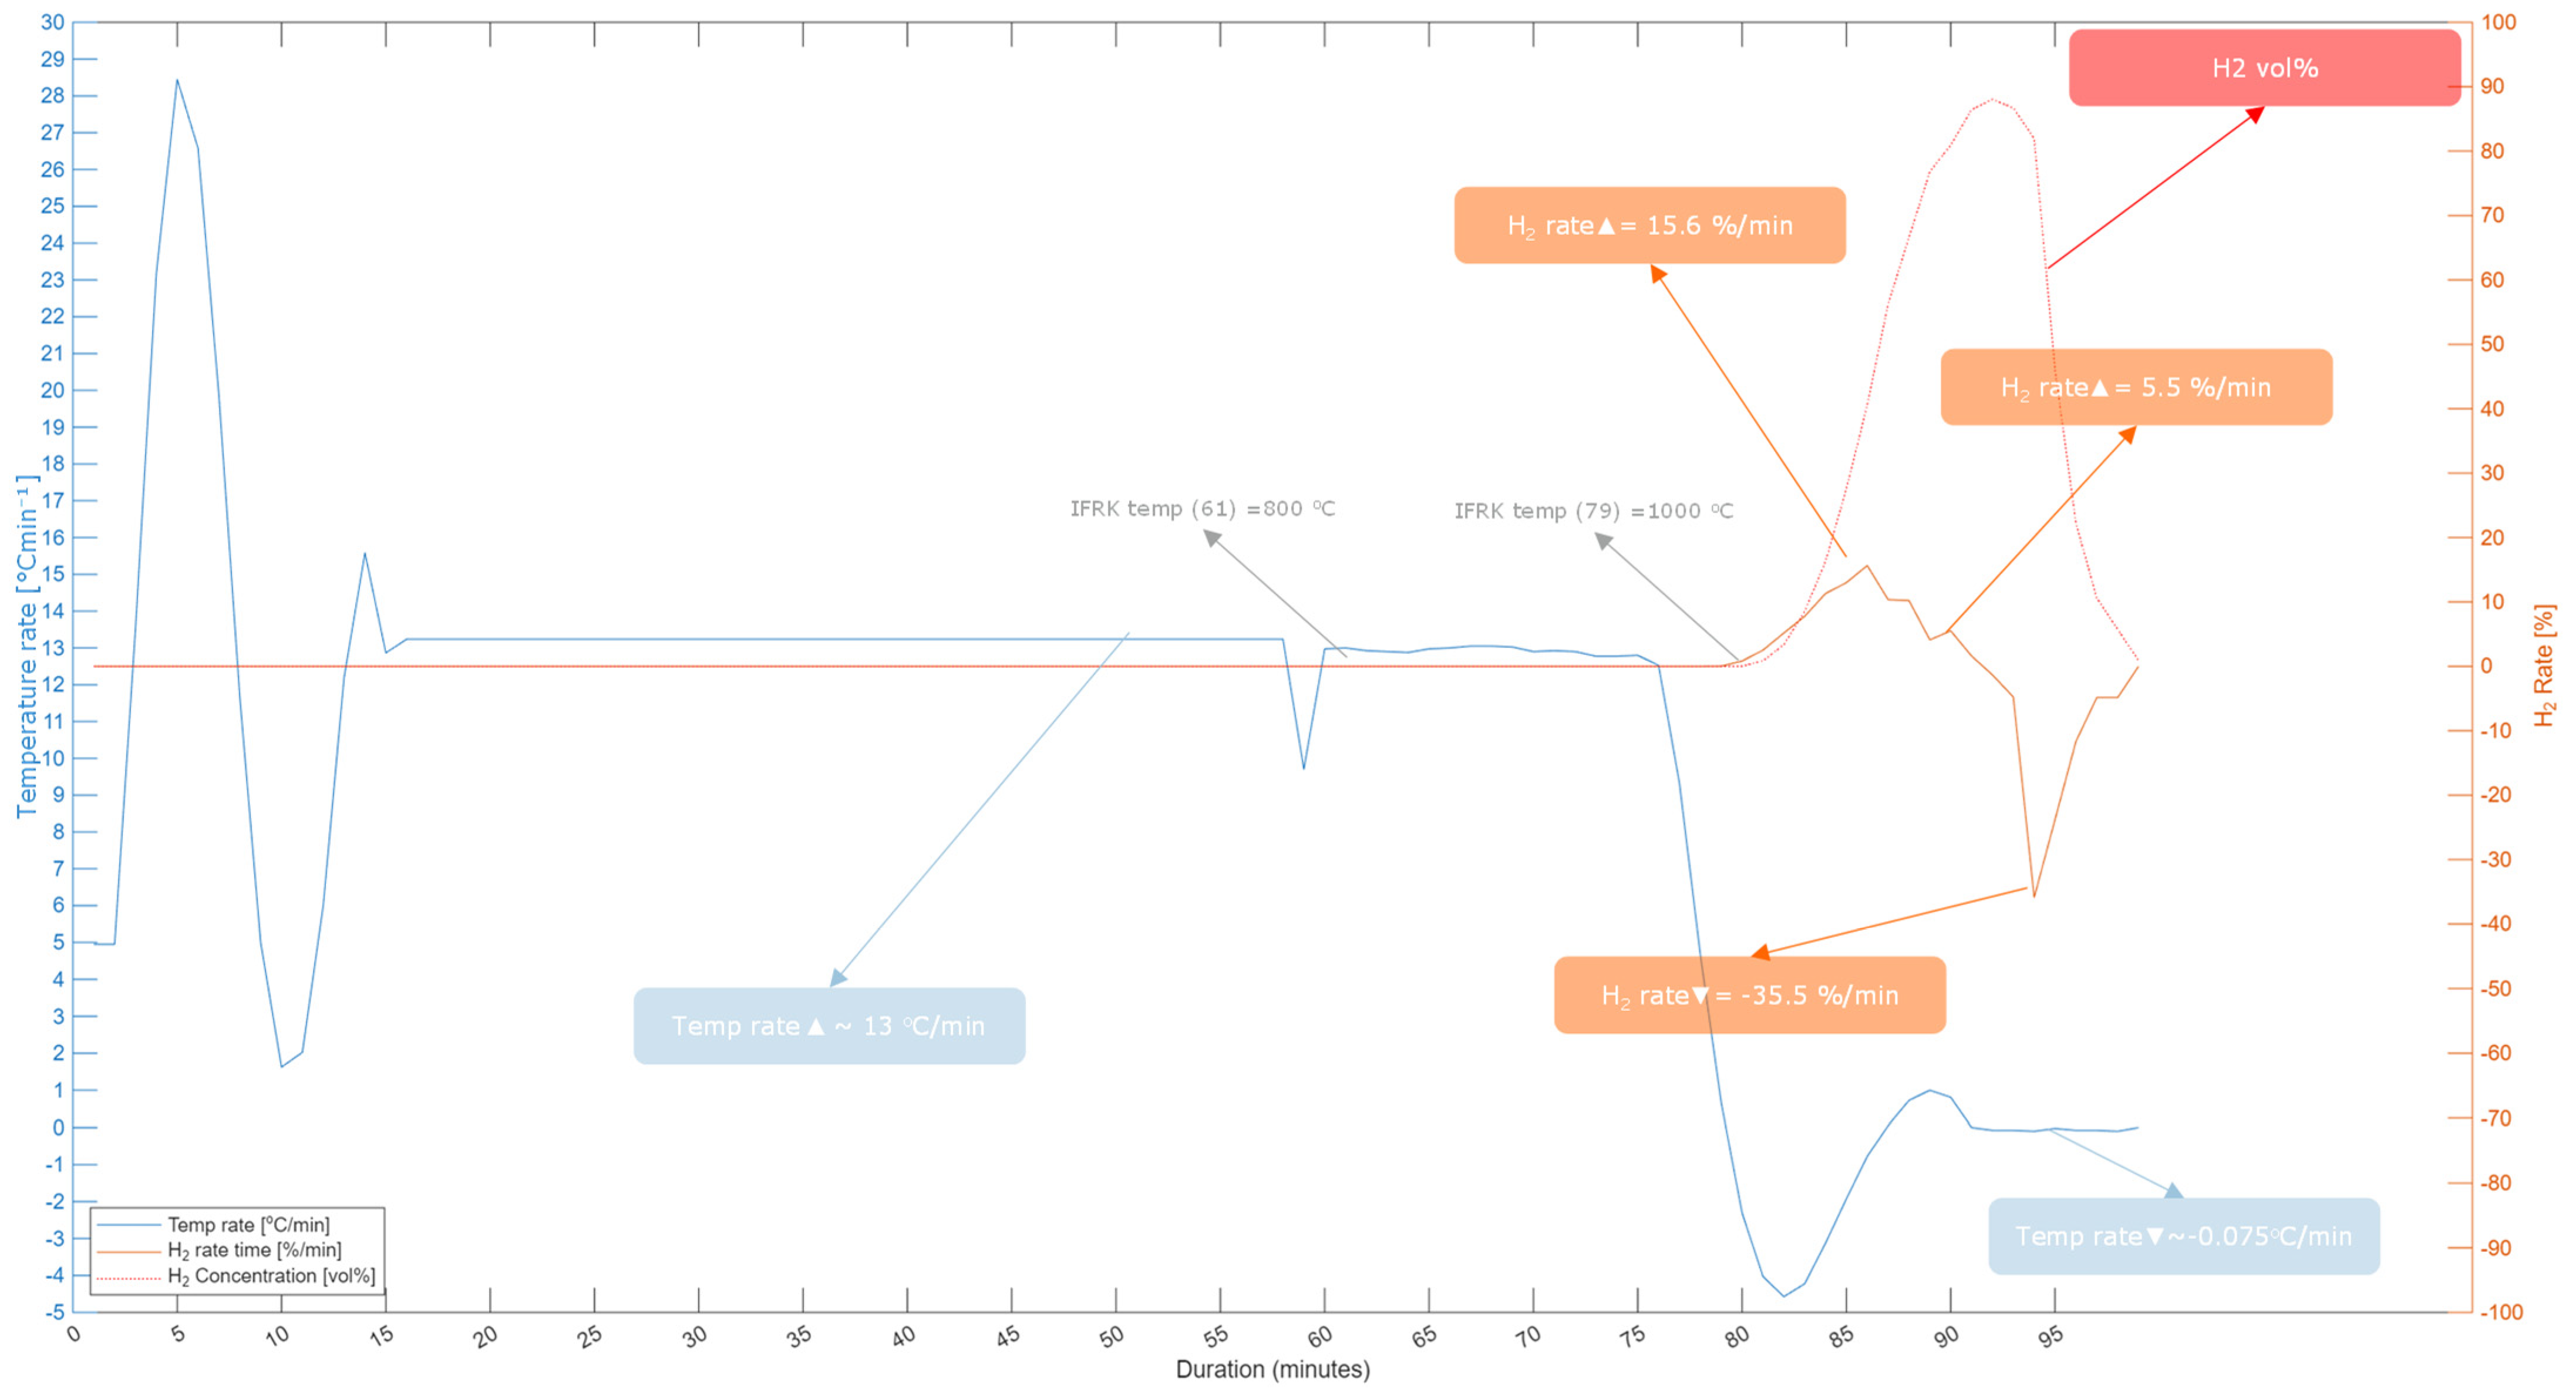
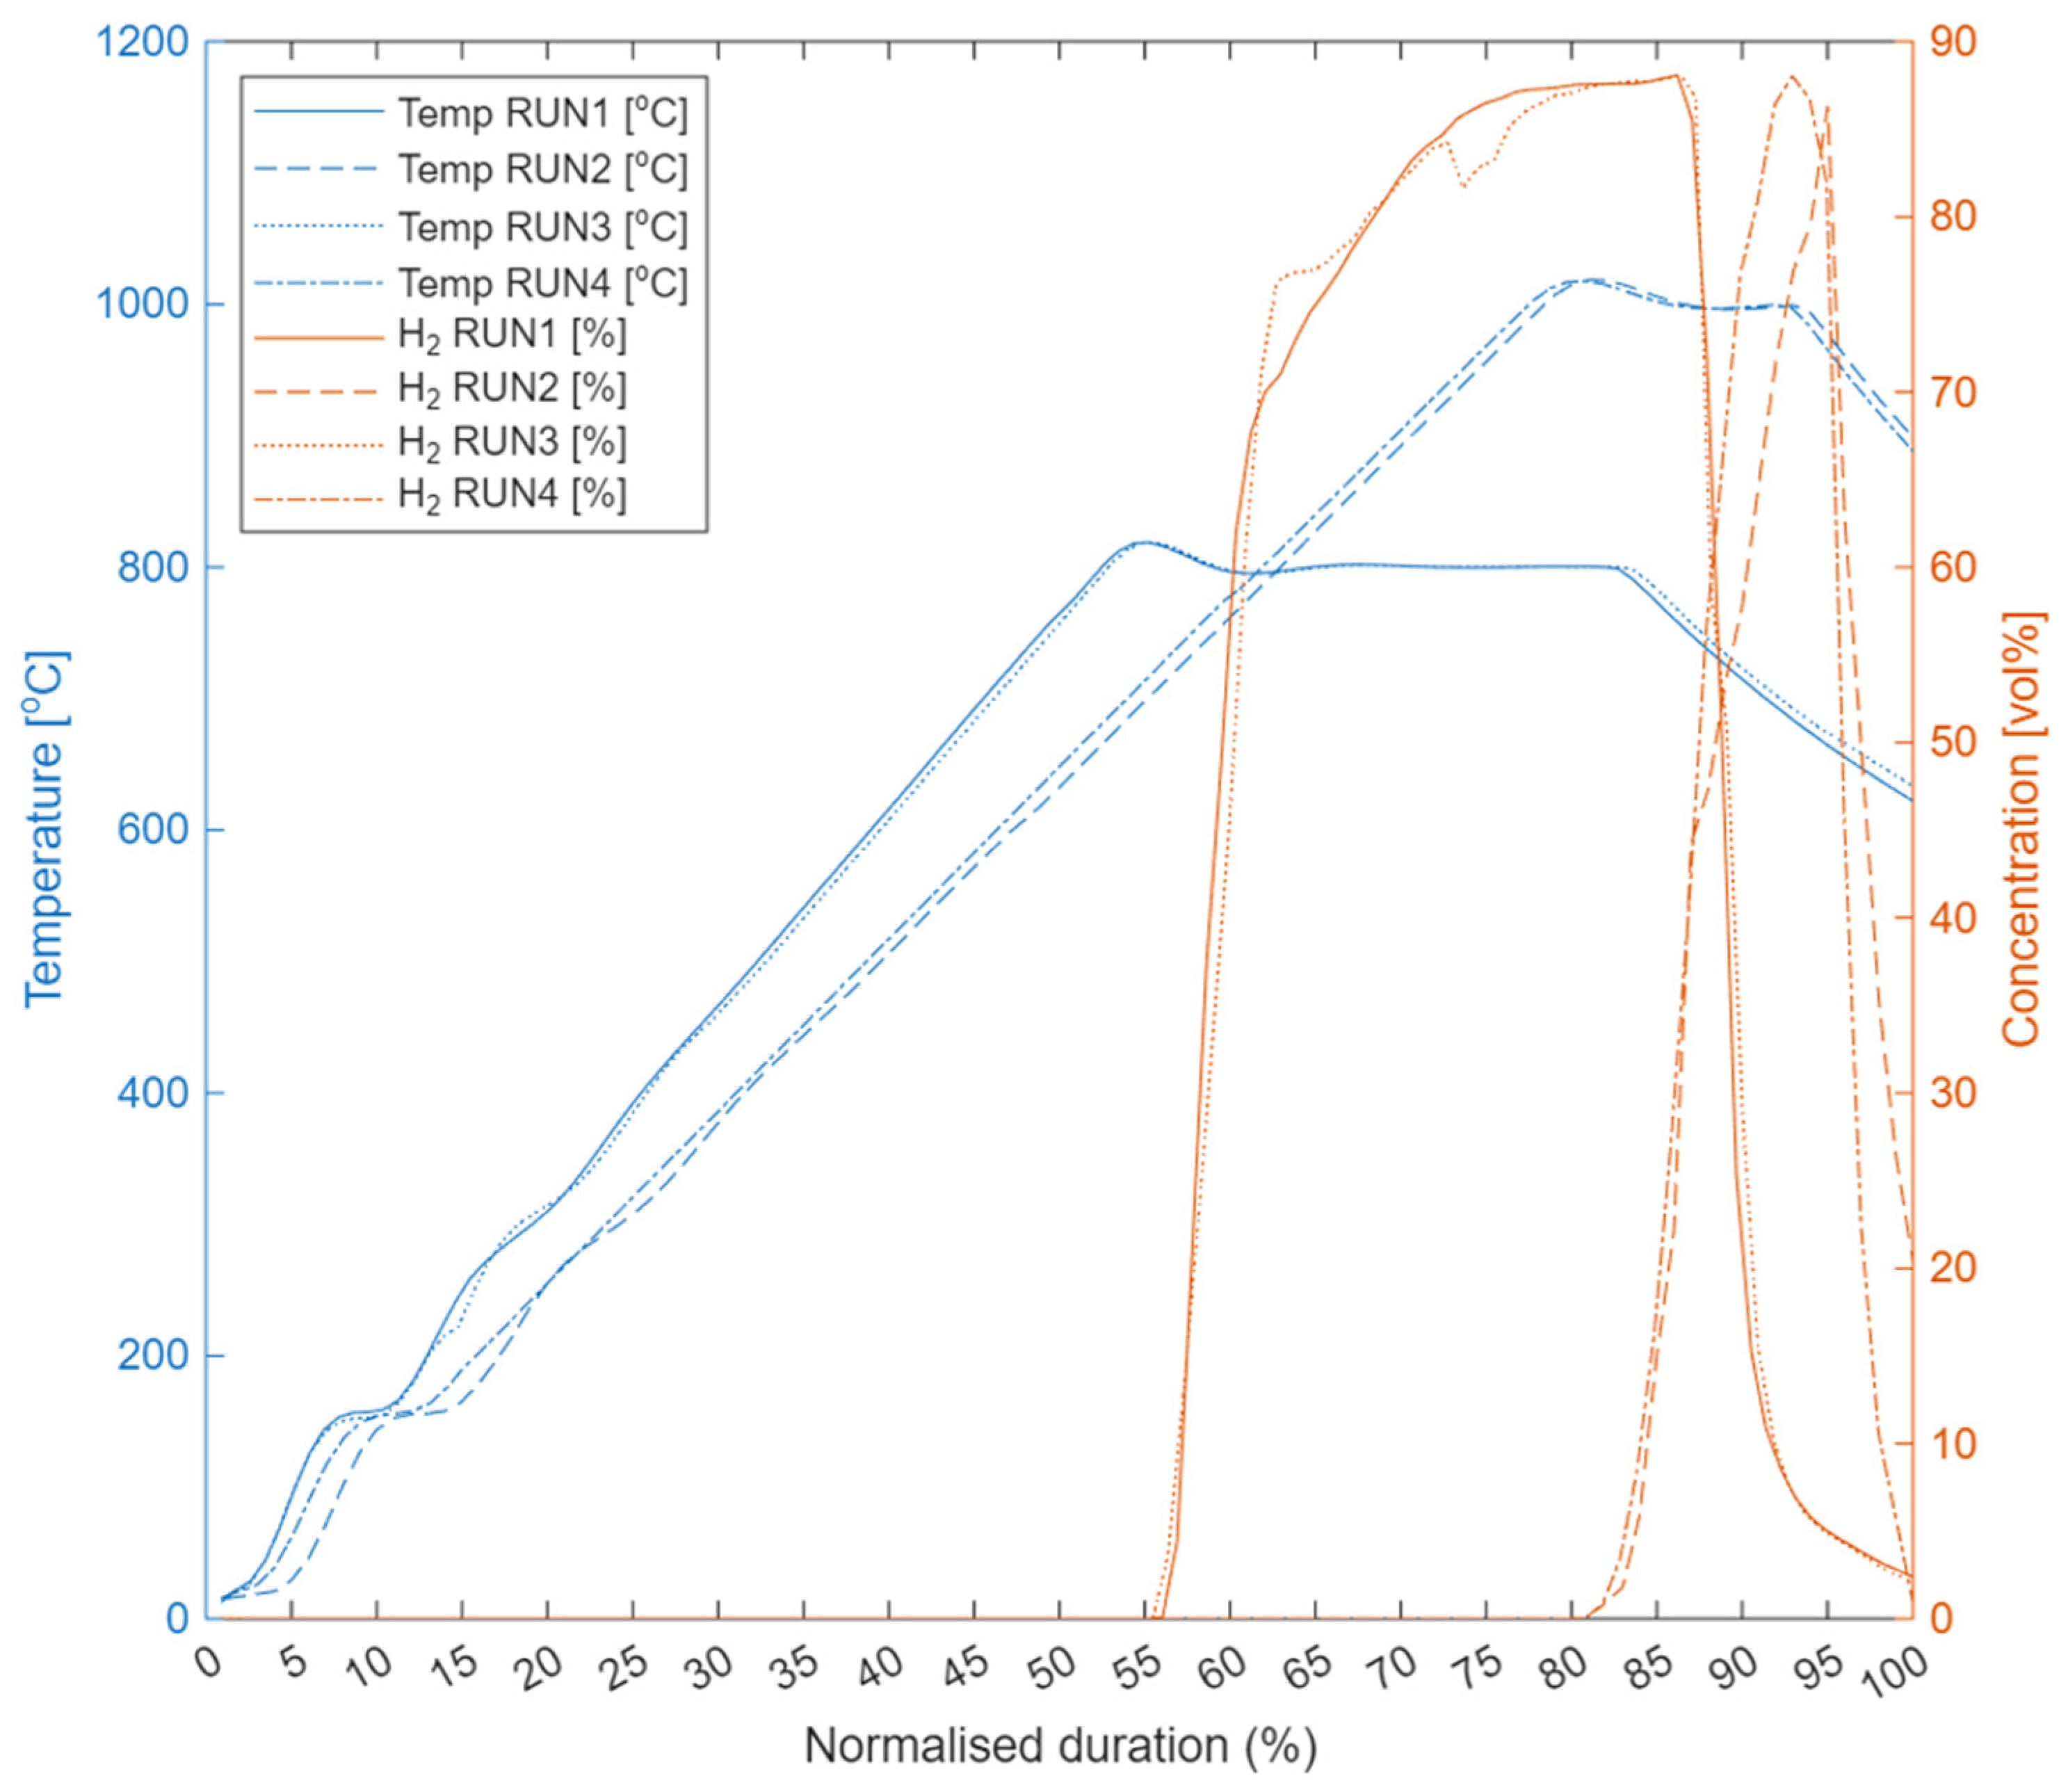
Heating was carried out in a flowing nitrogen atmosphere, which maintained inert conditions throughout the temperature ramp and prevented any reduction reactions before the desired operating temperature was attained. Once thermal equilibrium had been established, the nitrogen atmosphere was replaced by hydrogen, supplied at a nominal flow rate of approximately 0.55 standard L min−1—marking the beginning of the reduction stage. Two reduction temperatures were examined, namely, 800 °C and 1000 °C. To account for the expected increase in reaction rate at elevated temperatures, different hydrogen-treatment times were selected. The experiments at 800 °C were maintained under hydrogen for 35 min, whereas those conducted at 1000 °C were limited to 15 min. These conditions were selected to enable the evaluation of the temperature dependence of the reduction process while maintaining consistent operating conditions throughout the experimental campaign.
All experimental signals were recorded continuously through the laboratory data acquisition system. For quality assurance, only experiments with complete and uninterrupted datasets were used for the quantitative evaluation of temperature evolution, gas-flow behaviour and kinetic comparisons. During post-processing of the experimental data, a temporary communication interruption affected one reduction experiment (Reduction #4), resulting in incomplete controller and hydrogen-flow records. The interpolated values were used solely to preserve the continuity of the graphical presentation. They were not used in the calculation of reaction rates, kinetic parameters or any quantitative comparison. The interpretation of the reduction behaviour was based on the directly measured experimental data and independently supported by the physicochemical characterisation of the treated samples. Consequently, the interpolated intervals did not influence the interpretation of the overall reduction trend or the conclusions of this study.
2.5. Common Numerical Data-Processing Framework
A common numerical workflow was developed to process the measurements generated during the magnesite calcination and laterite hydrogen-reduction campaigns. Although the two processes were evaluated using different gas-phase indicators, both relied on the same underlying data architecture. Measurements from the IFRK controller, gas analyser and mass-flow controller were therefore imported, checked and aligned before the calculation of process-specific indicators. All numerical processing was performed in MATLAB R2026a [
32].
The workflow was designed to preserve the temporal sequence of each experiment while accounting for the different acquisition systems and measurement principles involved. The IFRK controller provided the thermal and operational history of the furnace, the gas analyser recorded the composition of the exhaust stream, and the mass-flow controller supplied information on the inlet-gas conditions. These datasets were combined into a common run-specific table indexed by elapsed experimental time. The processed datasets were subsequently used to calculate thermal, gas-phase and mass-balance indicators for the calcination and hydrogen-reduction experiments, while laboratory characterisation results (TG–DTA, XRD, SEM–EDS and mass measurements) were incorporated for the validation and interpretation of the calculated process indicators.
2.5.1. Data Acquisition
Three main data streams were used in the numerical analysis. The IFRK controller recorded the measurement timestamp, kiln temperature, setpoint temperature and heater output signal. These variables were used to reconstruct the thermal history of each run and to assess the response of the furnace to the programmed heating profile.
The outlet-gas composition was measured using the PGA. The recorded variables included timestamped concentrations of CO2, N2, H2 and O2 together with the analyser sample-flow measurement. CO2 was the principal gas-phase indicator during magnesite calcination, whereas H2 was monitored during the laterite-reduction experiments. Oxygen and nitrogen measurements were retained as a supporting indicator for the identification of air ingress, incomplete purging or disturbances in the gas line.
The mass-flow controller recorded the inlet volumetric flow rate, flow setpoint, gas temperature, absolute pressure and selected gas identity. These measurements were used to assess the stability of the nitrogen and hydrogen supply and, where the required flow information was available, to support the calculation of inlet-gas quantities.
2.5.2. Data Quality and Handling of Missing, Invalid, and Censored Measurements
Data-quality assessment was performed before the calculation of reaction indicators. Each dataset was checked for missing timestamps, duplicated observations, inconsistent units, abrupt discontinuities and values outside the expected operating range. Records associated with known calibration procedures or equipment checks were identified using the experimental log and the corresponding gas-concentration patterns.
To ensure continuity in the synchronised datasets, missing observations were treated using a linear imputation approach. Missing values were primarily associated with short communication interruptions between the monitoring platform and the connected instruments. Prior to imputation, all missing intervals were identified and documented for each variable and experimental run.
After synchronisation with the common time base, missing observations were filled using linear interpolation of neighbouring, no missing values (MATLAB\fillmissing with the ‘linear’ option). This approach estimates missing values from the nearest valid observations before and after the gap and is equivalent to linear interpolation between adjacent measured points. The implementation was applied only to short gaps bounded by valid observations and exhibiting a continuous local trend. Missing values in temperature, heater output, gas composition, and flow rate records were interpolated when the gap duration was sufficiently short and no known process disturbance occurred during the interval. Interpolation was not applied across periods corresponding to genuine operational transitions, such as gas switching, analyser calibration, intentional flow adjustments, pressure-release events or extended communication failures, since these events cannot be reliably represented by a linear trend. Long gaps and operationally significant interruptions were therefore retained as missing values and excluded from calculations requiring continuous data.
All preprocessing operations were documented by run. The data-quality record included the original observation count, missing values, duplicated records, interpolated points, calibration periods, censored measurements and excluded values. Data treatment was limited to the synchronisation, quality assurance and calculation of physically interpretable experimental indicators.
2.6. Numerical Indicators for Magnesite Calcination
The calcination case study was analysed by combining the solid mass balance, thermogravimetric measurements, particle-size distributions and the time-dependent CO2 concentration recorded at the IFRK outlet. The goal is to evaluate calcination performance from complementary perspectives, including chemical conversion, solid-product recovery, particle-size transformation and the temporal evolution of CO2 release during decomposition. The mass balance provides an estimate of the overall removal of volatile material and the recovery of the solid product. TG–DTA is used to determine the residual carbonate content of the treated material. Particle-size analysis describes the physical transformation of the feed during heating and rotation, while the gas-phase measurements identify the timing and duration of the principal decomposition period.
2.6.1. Solid Mass Balance
The solid mass balance is established from the initial feed mass and the mass of material recovered after each IFRK experiment. For pure MgCO
3, the theoretical fraction remaining after complete calcination (fMgO) is obtained from the molar masses of MgO and MgCO
3:
where the molar mass of magnesium oxide (MMgO) is 40.30 g/mol and the molar mass of magnesium carbonate (MMgCO
3) is 84.31 g/mol. Accordingly, complete decomposition of pure MgCO
3 theoretically leaves 0.478 of the initial carbonate mass as MgO, while approximately 52.2% is released as CO
2.
The theoretical final mass is estimated, also considering that the feed was not composed exclusively of MgCO
3, taking into account the non-reactive and secondary mineral fractions. If wMgCO
3 represents the mass fraction of MgCO
3 in the initial sample, the theoretical residual mass after complete decomposition (m
f,theor) can be calculated as
where m
0 is the initial sample mass and wMgCO
3 is the mass fraction of reactive MgCO
3. This simplified expression assumes that the non-MgCO
3 fraction remains in the solid phase. Where other carbonate phases contribute measurably to the thermal mass loss, the theoretical residual mass should instead be determined from the complete mineral-specific decomposition balance or directly from the initial TG–DTA mass-loss fraction. A TG-based formulation was therefore also used:
where Δmraw is the fractional mass loss of the untreated material over the carbonate-decomposition interval. This expression uses the experimentally measured volatile fraction and incorporates the combined contribution of all thermally decomposing phases detected under the selected TG–DTA conditions.
The mass-based calcination conversion was calculated as
where m
f is the measured recovered final mass. The quantities calculated for each experiment were organised according to
Table 1.
2.6.2. TG-Based Residual Carbonate and Calcination Degree
Thermogravimetric and differential thermal analysis was used to establish the decomposition behaviour of the untreated magnesite and to determine whether carbonate remained after IFRK processing. The untreated sample was first analysed to identify the decomposition interval and the corresponding total mass loss. Treated samples were analysed under the same or directly comparable conditions so that the remaining decomposition-related mass loss could be expressed relative to that of the raw feed.
The TG–DTA was applied to both untreated and IFRK-treated magnesite samples. The resulting thermograms and differential thermal analysis curves are presented and discussed in
Section 3, where the residual carbonate content and calcination degree are evaluated based on the measured mass-loss behaviour and thermal events.
2.6.3. Particle-Size Analysis
Particle-size analysis was performed before and after calcination to determine whether thermal decomposition and rotary motion altered the physical size distribution of the material. Coarser particles were analysed by sieving, while the finer fraction was analysed using laser-diffraction measurements where applicable. The analysis focused on changes in the cumulative passing curves and on characteristic particle-size indicators derived from the measured distributions.
For each sieve size, the mass fraction retained was calculated as
where w
i is the mass fraction retained in size interval i and m
i is the mass retained in size interval i.
Characteristic particle sizes were determined from the cumulative passing curve. D50 represents the particle size below which 50% of the sample mass is found, while D80 corresponds to the size below which 80% of the sample mass passes. Where the required passing percentage occurred between two measured sieve sizes, logarithmic interpolation was used because particle-size distributions are conventionally represented on a logarithmic size axis. Consequently, the particle-size results describe the net physical transformation produced by the complete treatment procedure and not exclusively the chemical decomposition mechanism.
2.6.4. Comparative Analysis of Operating Variables
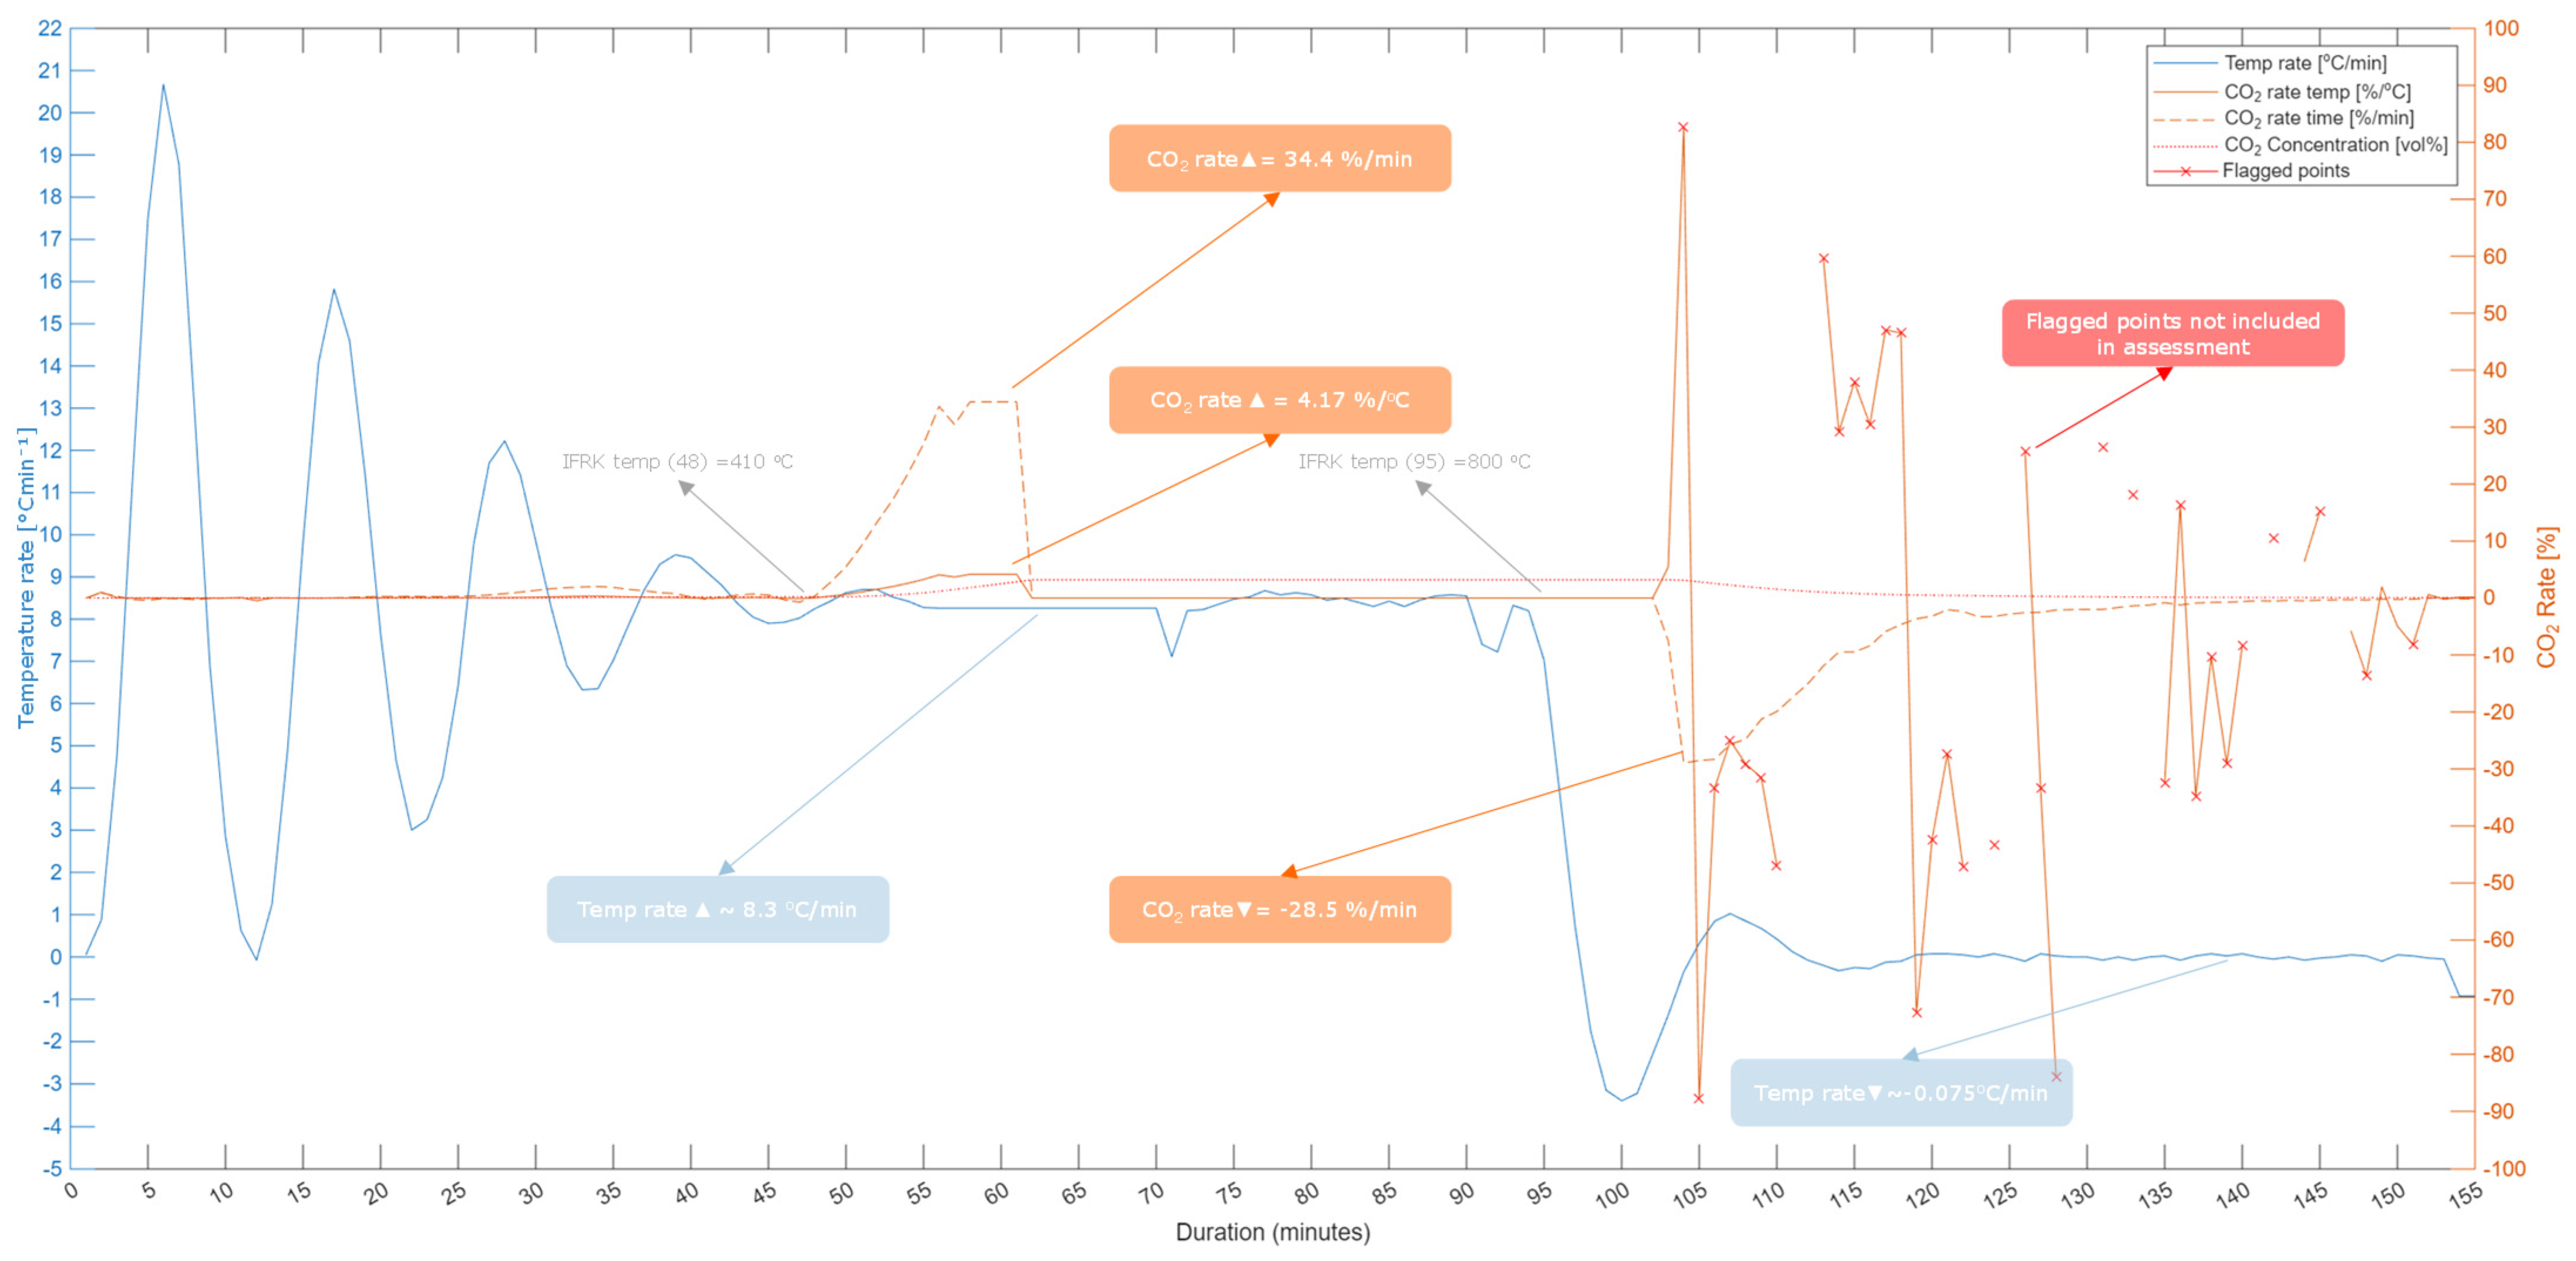
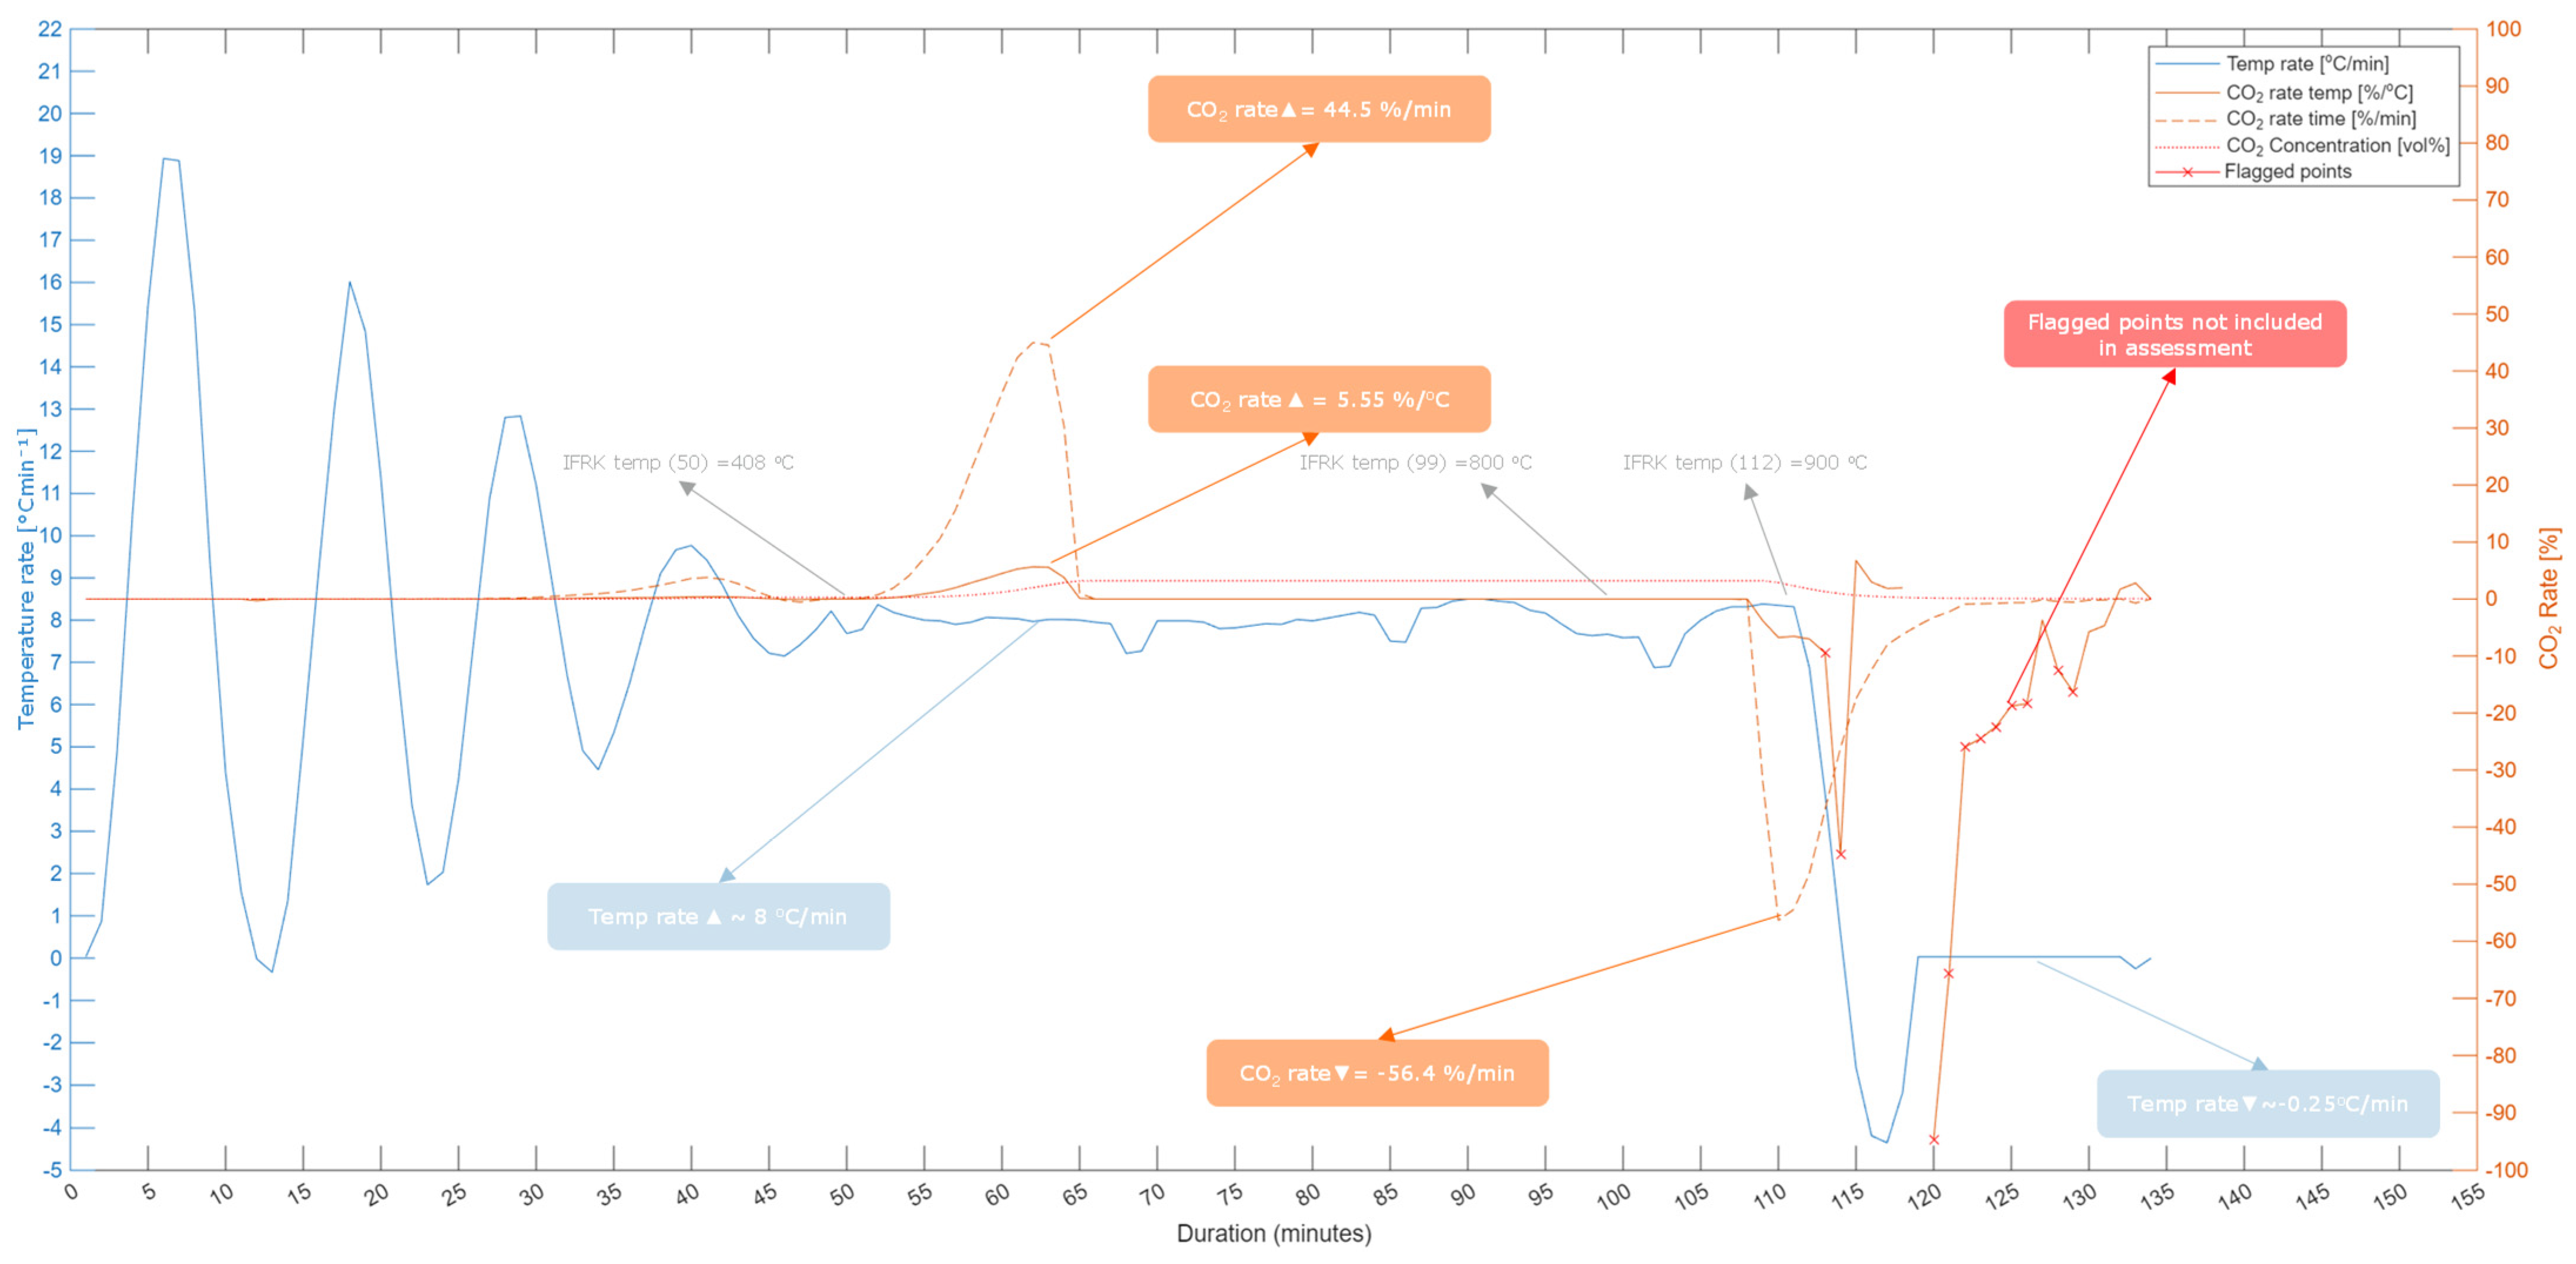
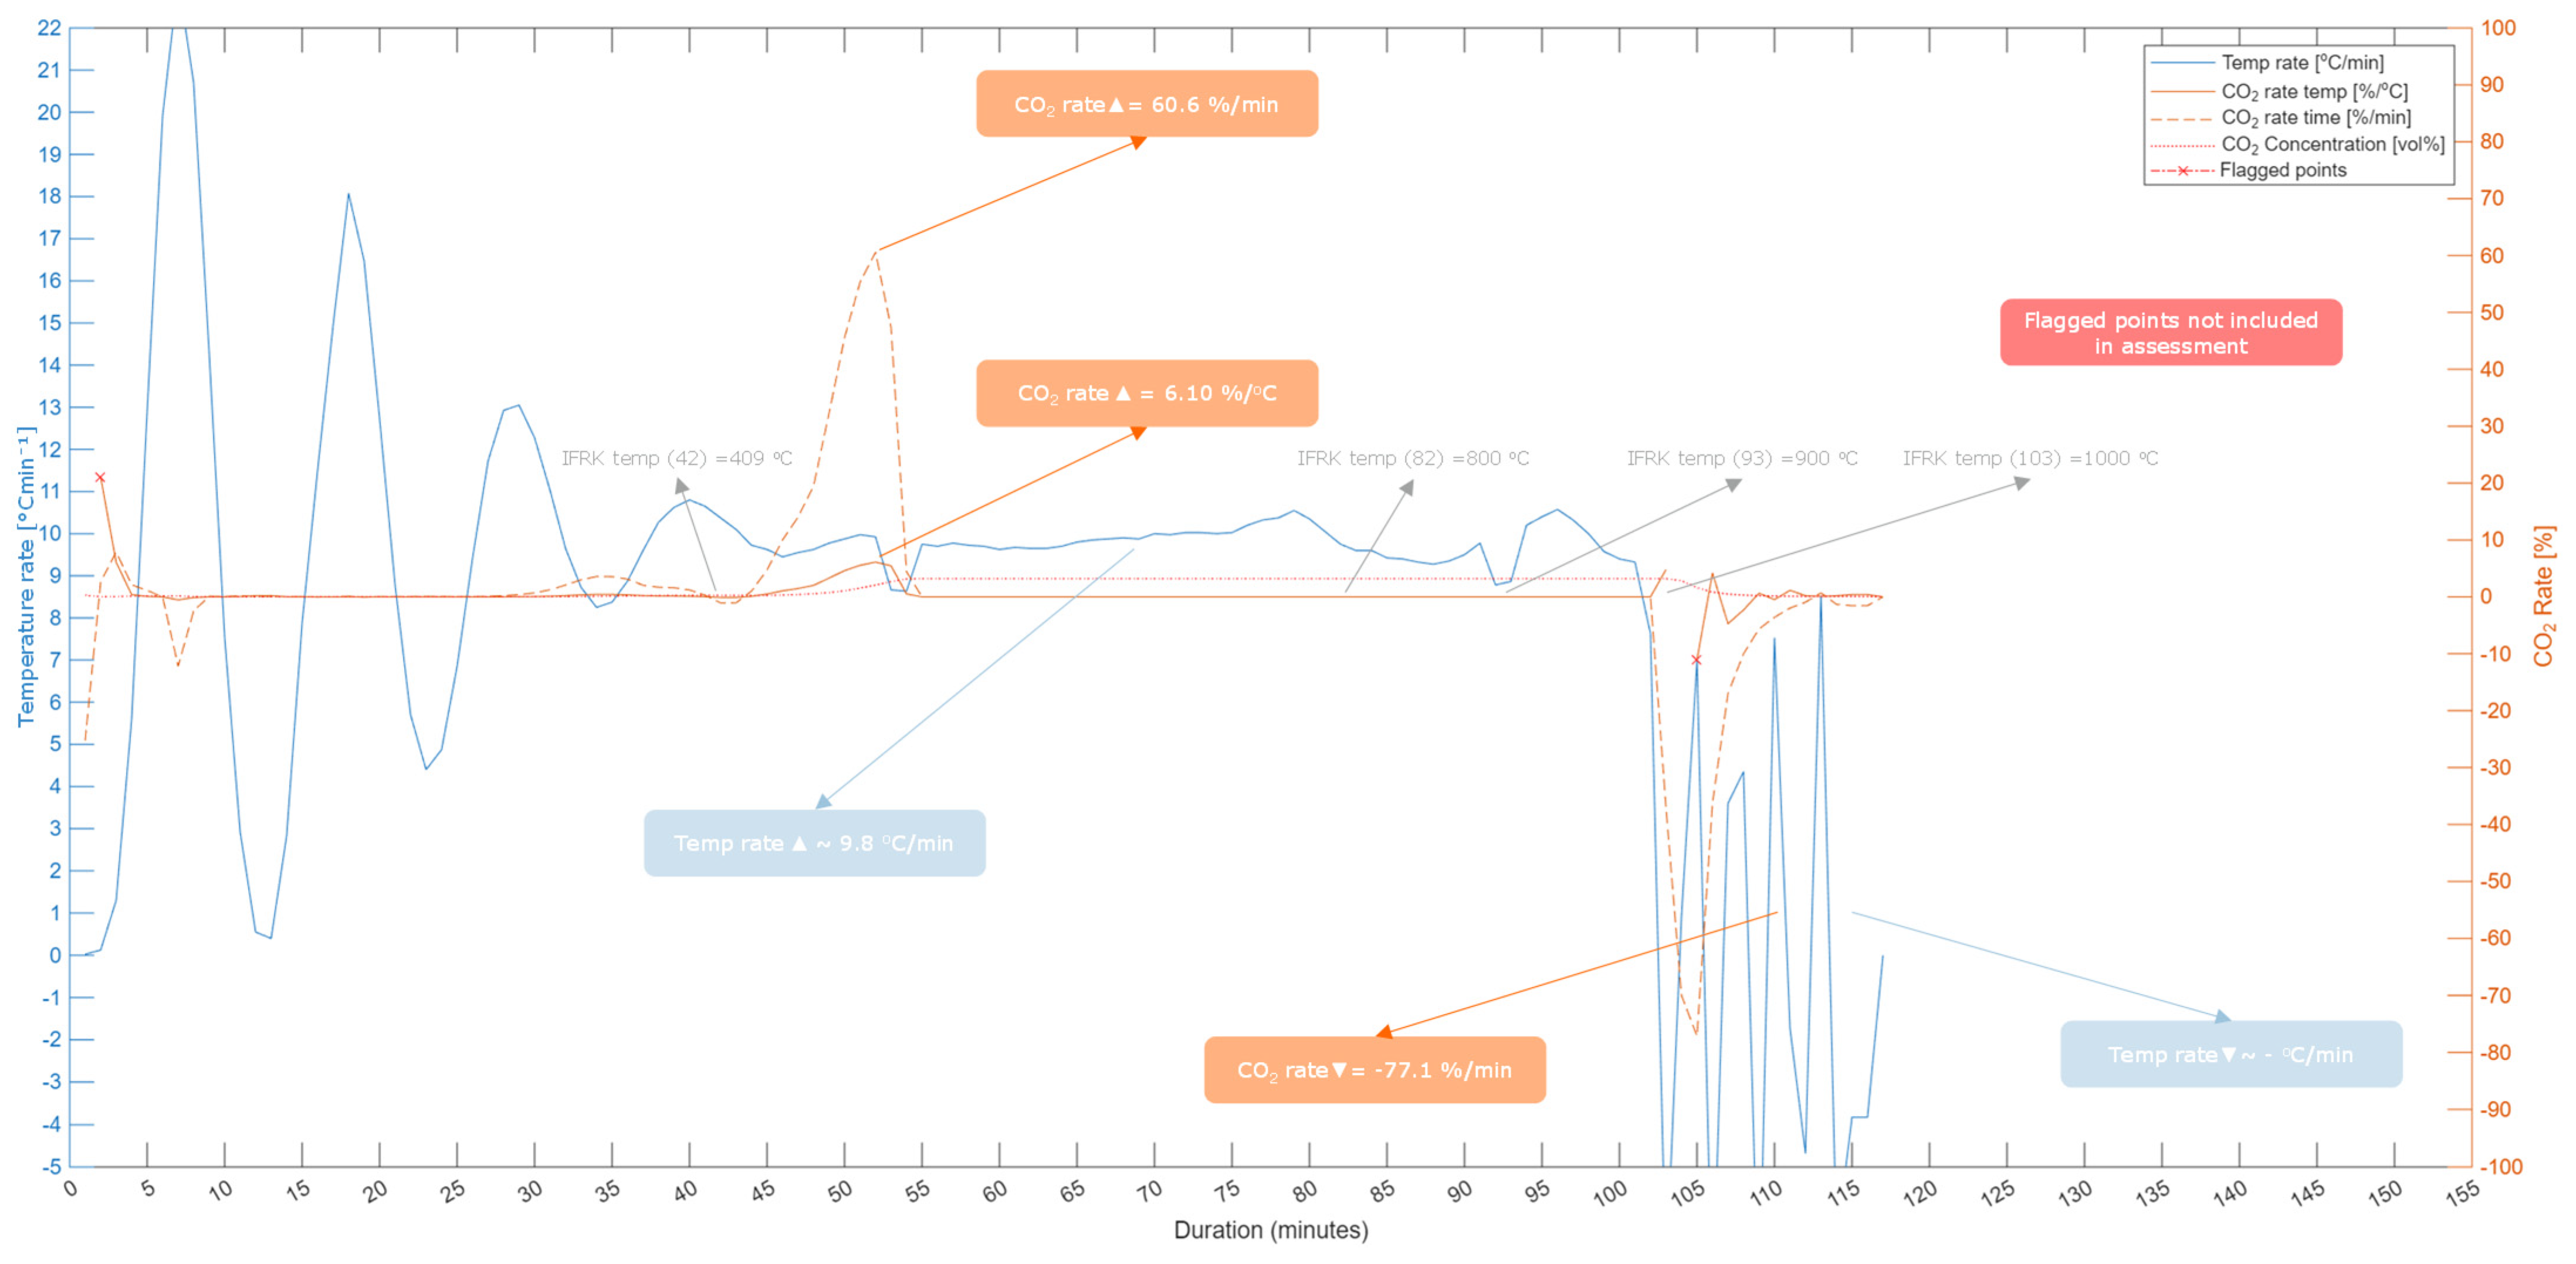
The magnesite experiments were conducted at target temperatures of 800, 900 and 1000 °C. The principal comparison, therefore, concerned the influence of the maximum temperature on the thermal history, CO2 response, processing duration and final solid condition. However, the experiments did not constitute a fully balanced design because the initial sample mass and the actual heating rate were not identical in all runs. The measured heating rates ranged approximately from 8.0 to 9.8 °C min−1. Direct comparison of raw gas-concentration values could therefore reflect differences in sample inventory and operating history in addition to the target temperature.
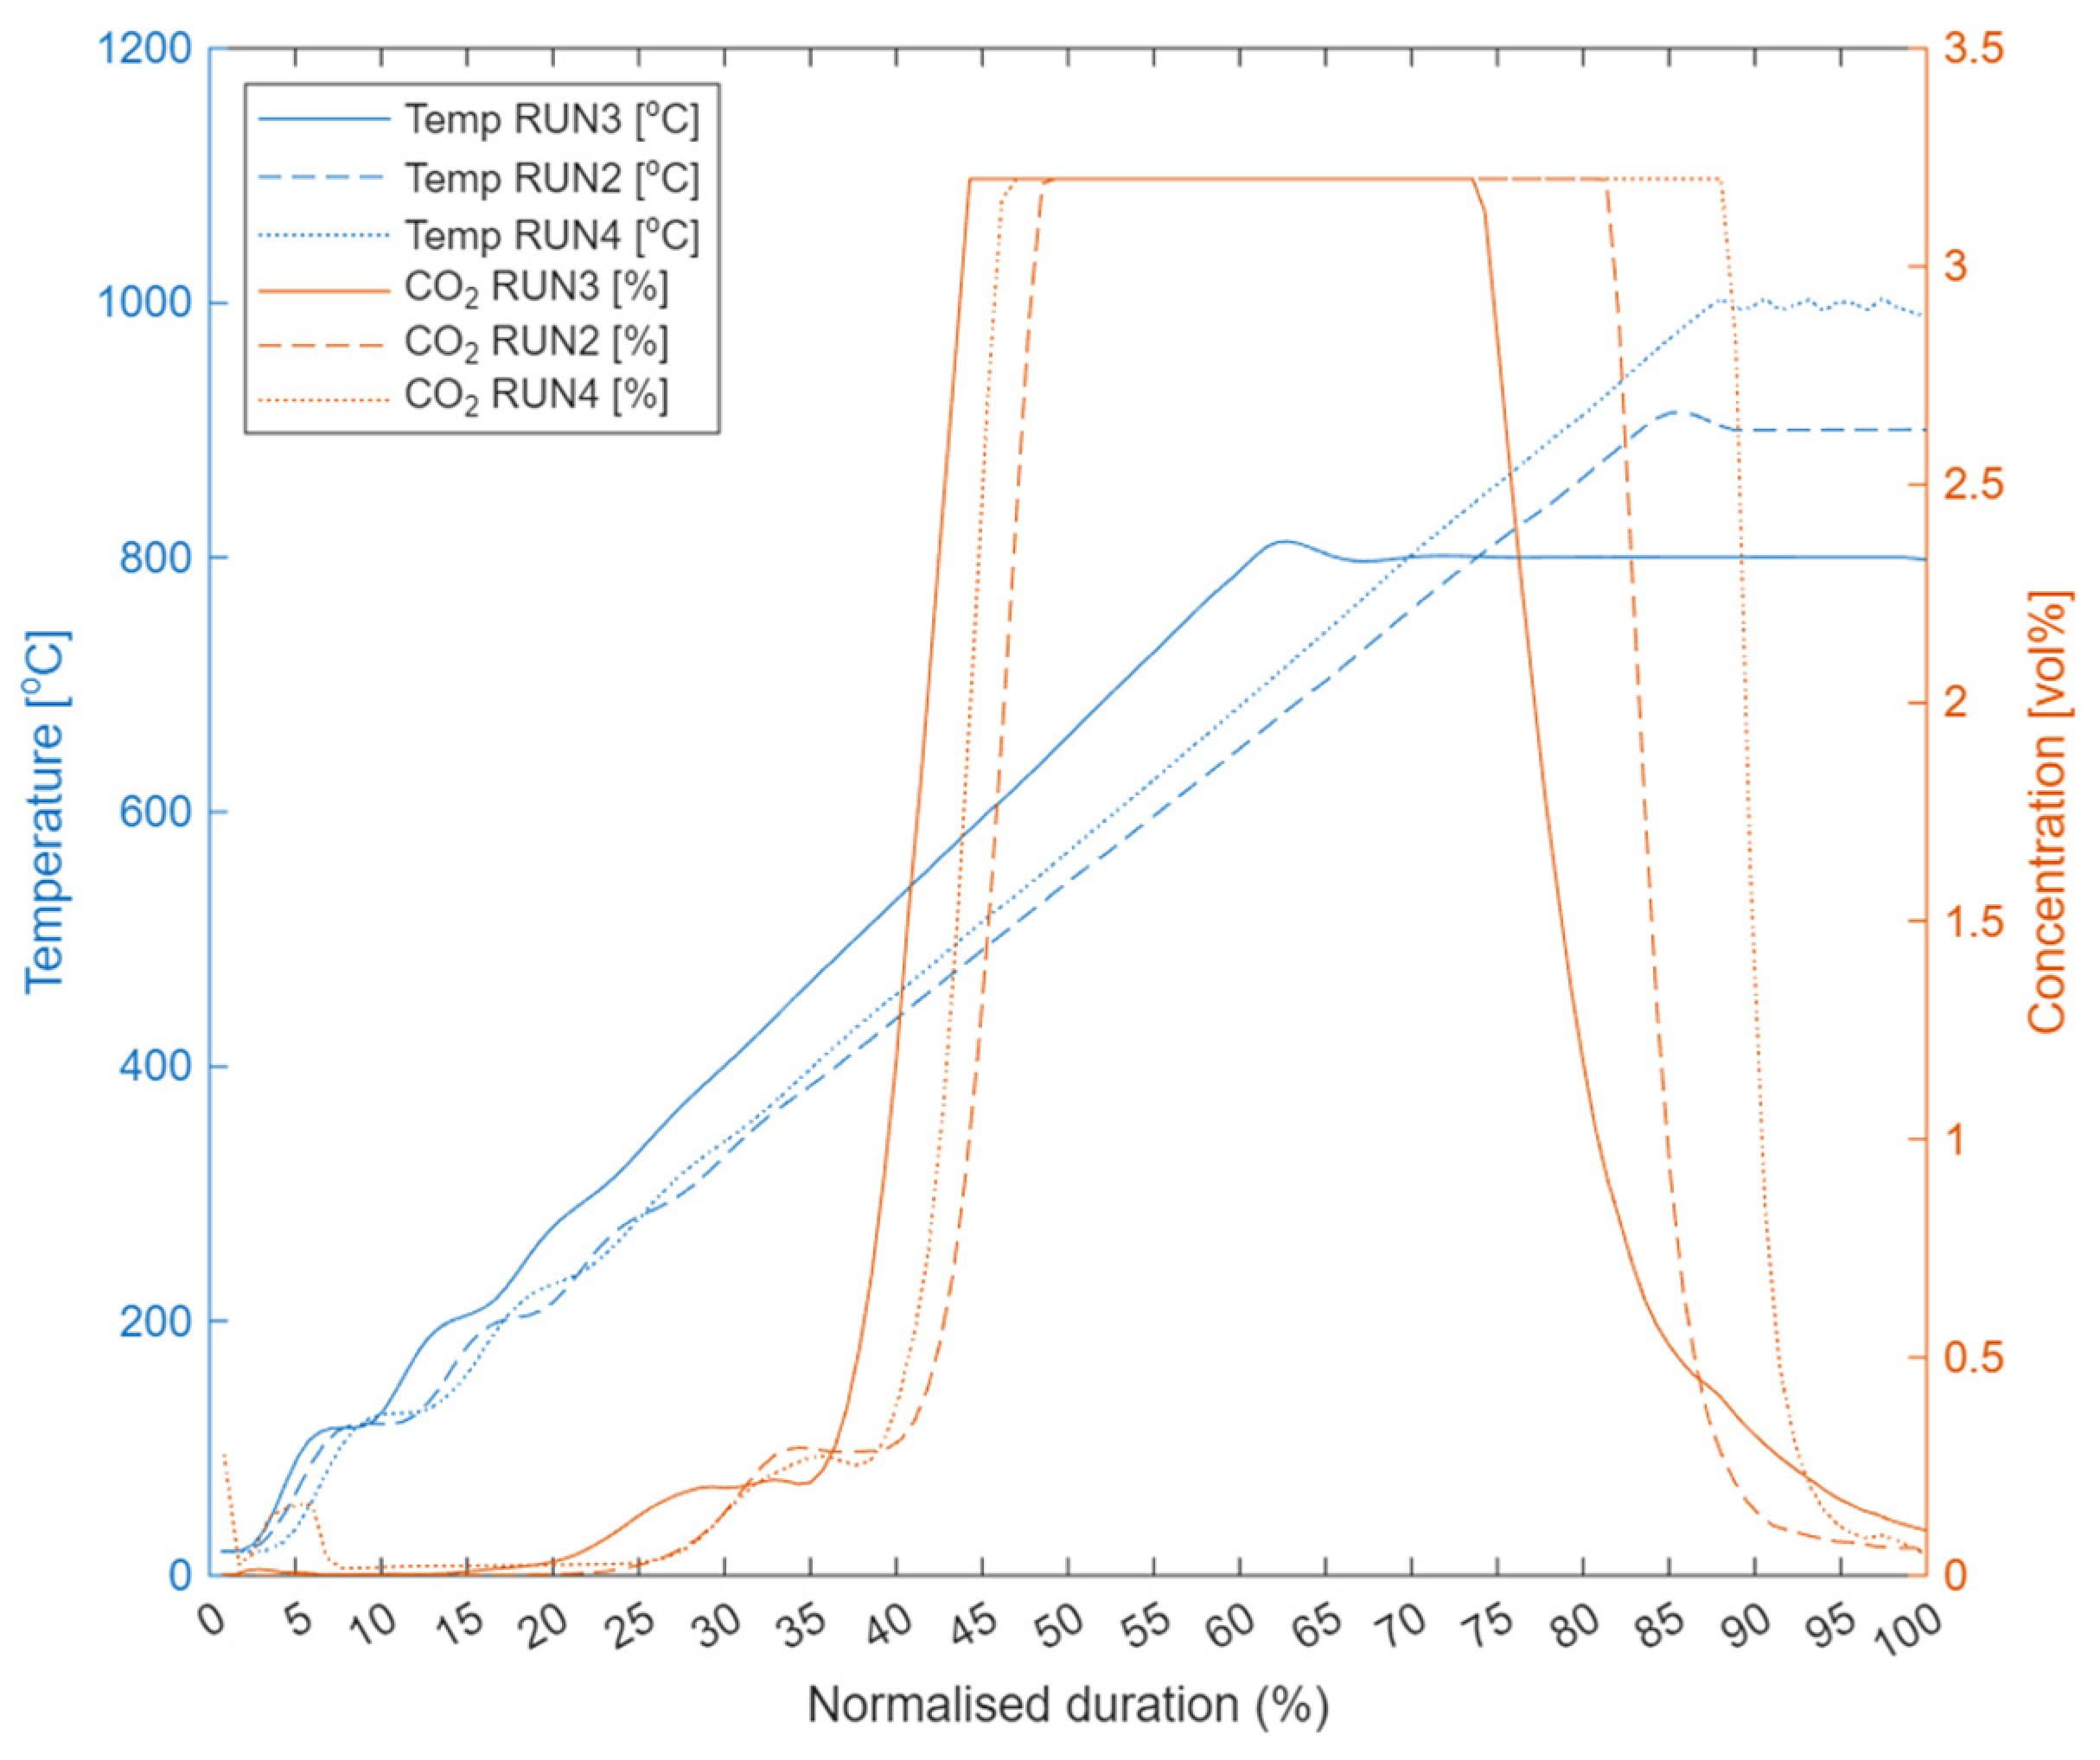
The comparative evaluation was based on analyses performed using the synchronised temperature and CO2 datasets. The temperature profiles over time were compared using the actual average heating rate, the time required to reach the target temperature, and the total run duration. In addition, the synchronised CO2 measurements were evaluated in terms of the onset of detectable CO2 evolution, the duration of the main CO2-evolution period, and the timing of CO2 depletion.
The influence of target temperature was evaluated using the measured mass loss, TG-based calcination degree, product recovery, particle-size distributions and the temporal characteristics of the CO2 signal. Because only one run was available at each main condition and the sample masses differed, the comparison was treated as exploratory. The observed differences were therefore interpreted as experimental trends rather than statistically replicated temperature effects.
Particle-size analysis was used to compare the feed material and the calcined product and to assess the physical transformation associated with calcination and kiln rotation.
2.6.5. Gas-Control Performance and CO2-Evolution Indicators
The outlet CO2 concentration was used to identify the temporal development of the calcination reaction. Since the analyser measured gas concentration, the primary indicators were formulated as concentration-based temporal metrics. These indicators were used to compare the timing and relative shape of the CO2 response between experiments.
The beginning and end of the principal CO
2-evolution interval were identified from the sustained rise above baseline and the subsequent return toward the baseline range. The duration of the interval was calculated as
where t
end was defined as the first point after the main release period at which the concentration fell below the selected endpoint threshold and remained below it for a specified period.
The end of the CO
2-evolution period was identified using a concentration threshold of approximately 0.6 vol.% CO
2. This threshold was established after reviewing the CO
2 profiles from all calcination experiments and was found to consistently indicate the point at which the main decomposition stage had essentially finished and the outlet gas entered the final depletion phase. The selected value also remained comfortably above the normal baseline fluctuations of the gas analyser, minimising the influence of measurement noise while providing a consistent basis for comparing the different experimental conditions. The concentration-rise slope was calculated as the change in CO
2 concentration divided by the corresponding change in time. The concentration-rise slope (rCO
2) was calculated as
where ΔyCO
2 is the change in measured CO
2 concentration and Δt is the corresponding change in time.
Where the temperature dependence of the signal was examined, the local concentration change with temperature was expressed as the CO
2 concentration change per unit temperature (rCO
2,T):
Both slopes were calculated only from valid, unsaturated measurements. Data points at the analyser ceiling were treated as censored observations and were excluded from numerical differentiation. The combination of these indicators allowed the calcination experiments to be examined at three complementary levels: chemical conversion of the solid, physical recovery and fragmentation of the product, and temporal development of the gaseous decomposition response.
2.7. Numerical Indicators for Laterite Hydrogen Reduction
The hydrogen-reduction experiments were evaluated by combining the recorded IFRK temperature and outlet-gas measurements with the initial and final sample masses and the post-treatment mineralogical and microstructural analyses. The purpose of this combined approach was to assess the progress of reduction from both the gas and solid phases. The H2 concentration profiles were used to examine the response of the reactor after the transition from nitrogen to hydrogen, while the mass measurements provided an overall indication of the material changes occurring during heating and reduction. XRD and SEM–EDS analyses were subsequently used to identify the reduction products and to examine the distribution of iron-, nickel-, cobalt- and oxygen-containing phases in the treated samples.
2.7.1. Solid Mass Balance
The solid mass balance was determined from the mass of the laterite sample before and after each reduction experiment. The measured mass loss (Δm) was calculated as
where m
0 is the initial sample mass and m
f is the final sample mass.
The corresponding relative mass loss was calculated as
where Δm
rel is the relative mass loss expressed as a percentage of the initial sample mass. This quantity represents the total mass removed during the complete thermal treatment. It includes the heating stage under nitrogen, the hydrogen-reduction stage and the subsequent cooling and product-recovery operations.
The measured mass loss could not be treated as a direct reduction degree because several processes may contribute to the decrease in sample mass. These include the removal of free and structurally bound water, the dihydroxylation of hydrated silicate phases, the decomposition of carbonate phases, the removal of oxygen from iron and nickel oxides, and the physical loss of fine particles during handling or recovery. Furthermore, the raw laterite had a measured loss on ignition of 5.39 wt.%, indicating that a proportion of the total mass loss could occur independently of hydrogen reduction. The initial mass, final mass, absolute mass loss and relative mass loss were therefore reported together with the gas-phase and mineralogical results.
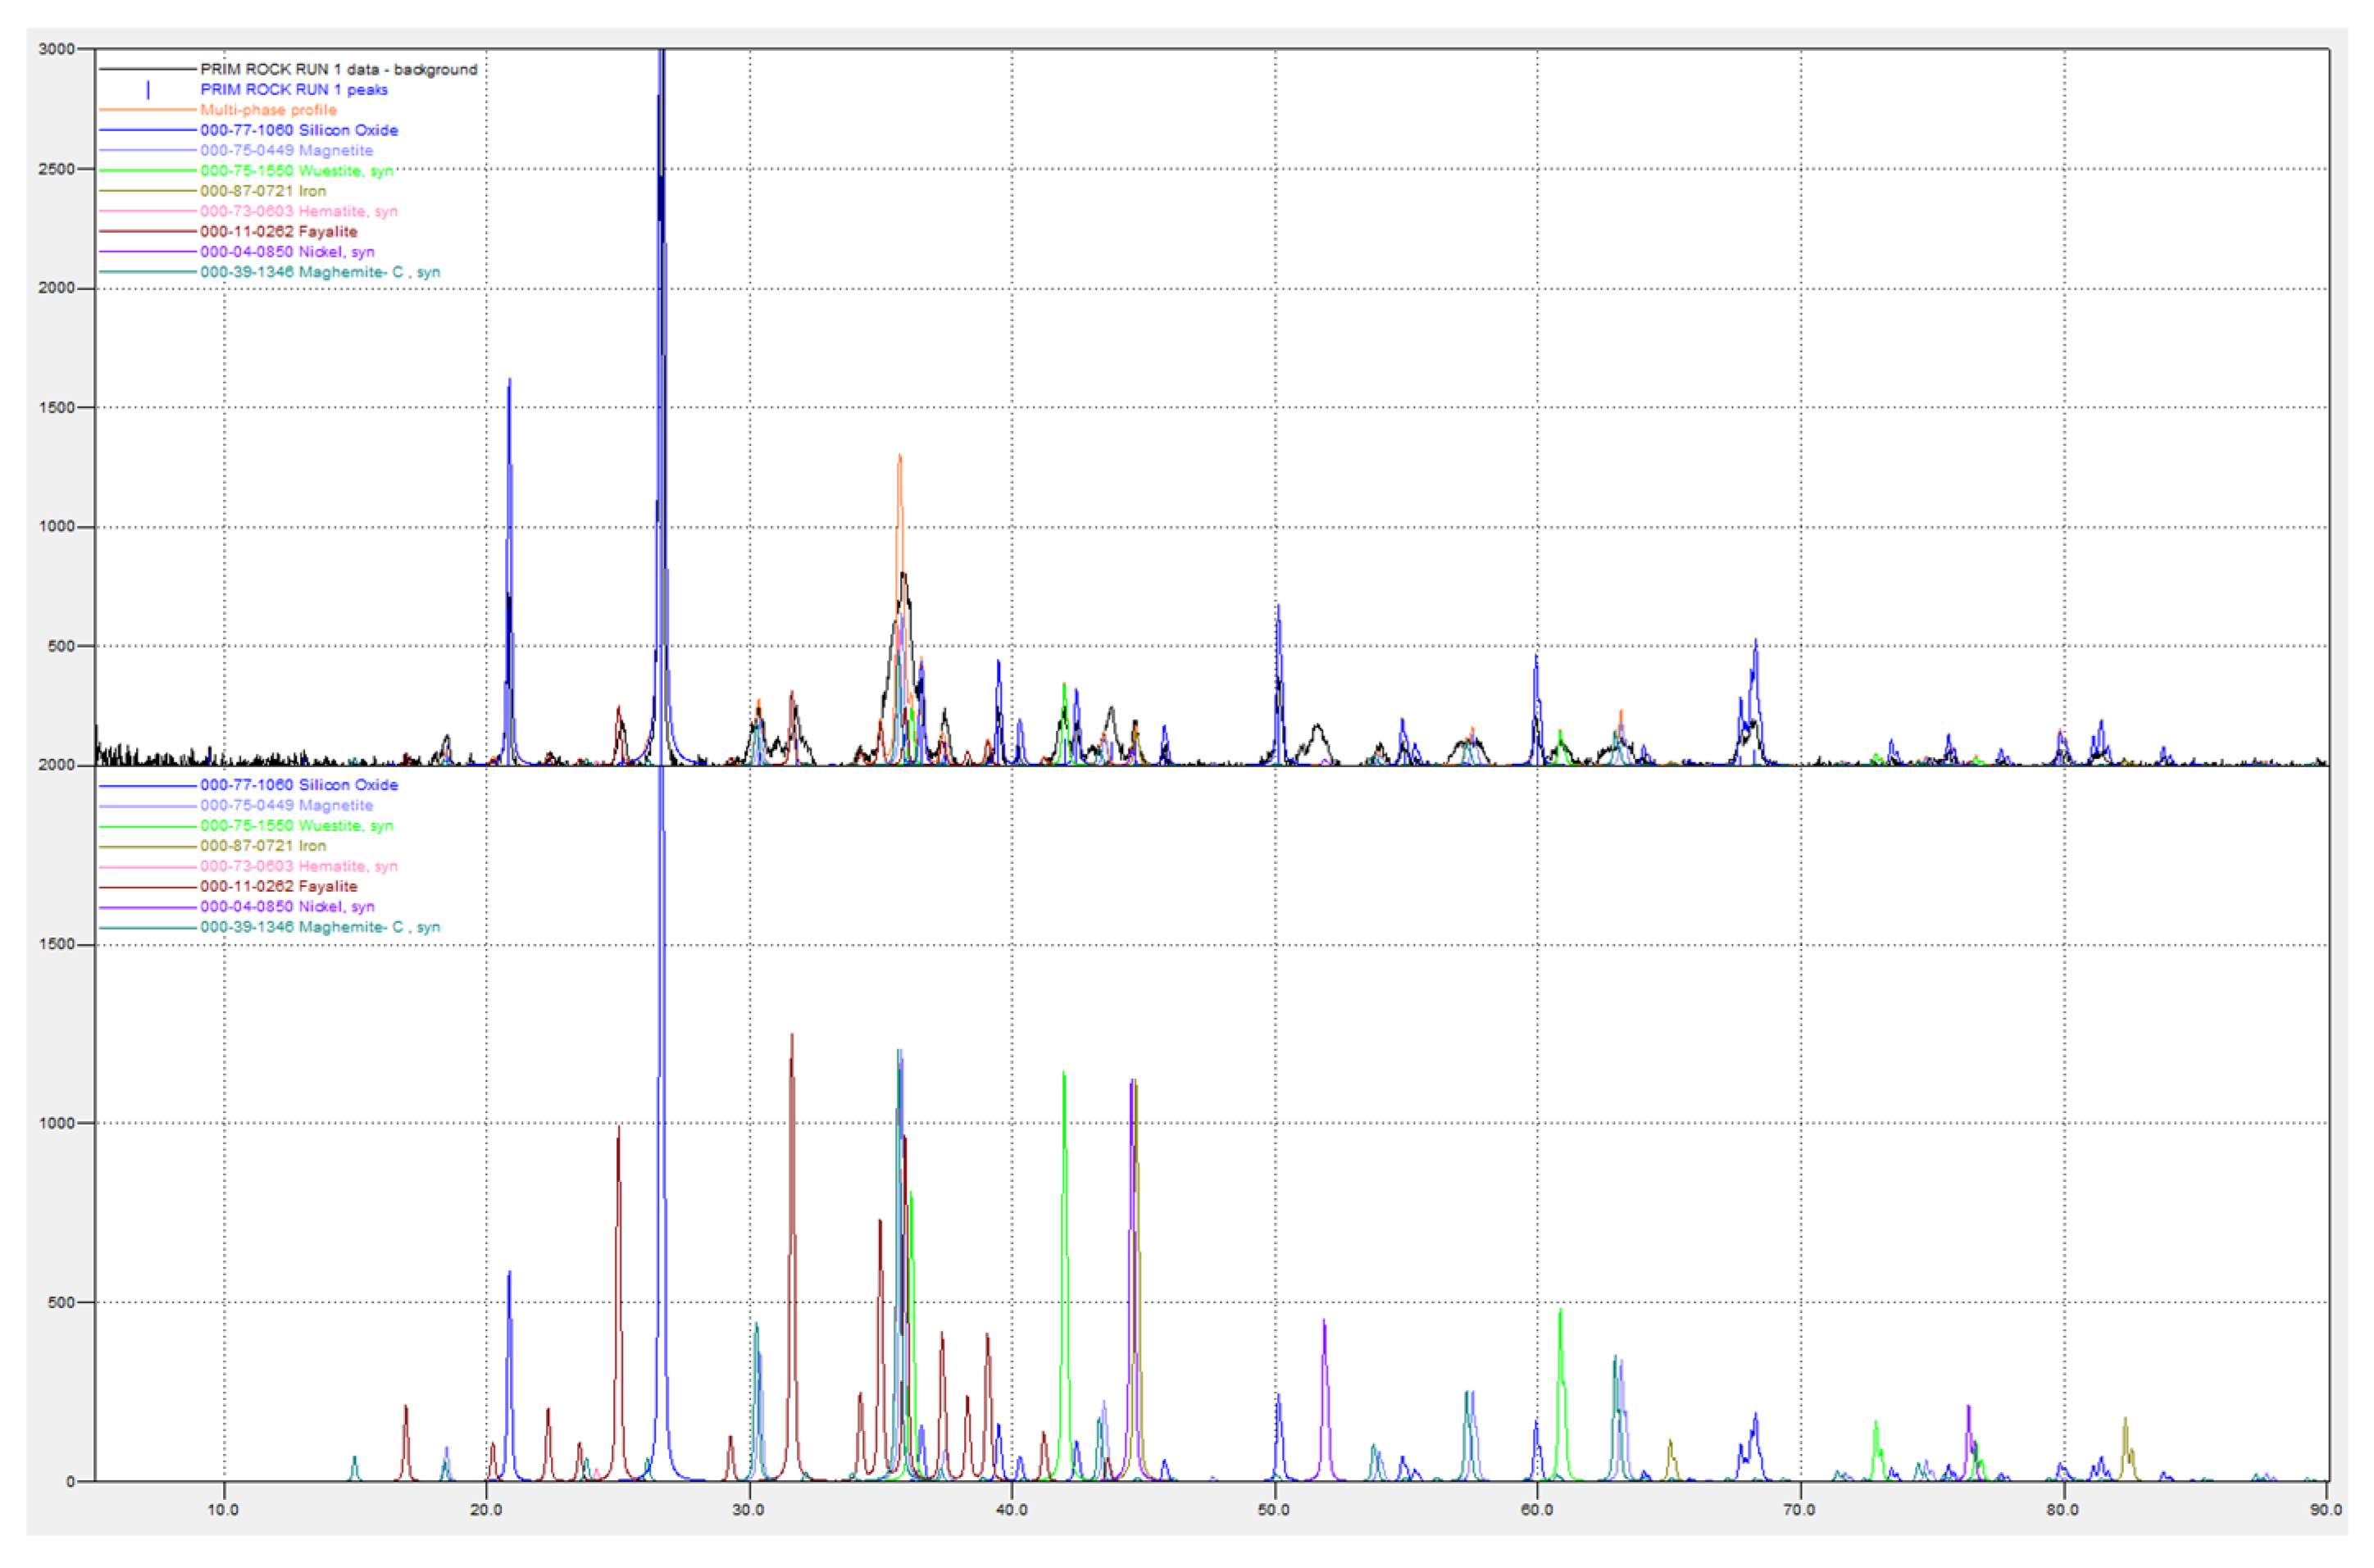
2.7.2. XRD Analysis of the Reduction Products
XRD was used to identify the mineral phases present after hydrogen treatment and to evaluate the progression of the reduction sequence. The untreated laterite was used as the mineralogical reference. Its principal identified phases included hematite, quartz and calcium carbonate, with possible minor contributions from clinochrysotile and magnesium chromite.
The XRD patterns of the treated samples were examined for changes in the iron-bearing phases. Particular attention was given to the reduction or disappearance of hematite reflections, the formation of magnetite and wüstite, the presence of fayalite or other iron-bearing silicate phases, the appearance of metallic iron and metallic nickel, and the changes associated with treatment temperature. The reduction pathway was interpreted according to the progressive transformation of iron oxides:
The identification of intermediate phases was used as evidence of partial reduction, whereas the presence of metallic iron and the disappearance of hematite were used as evidence of more advanced reduction. Since laterite contains several overlapping crystalline phases, phase identification was based on the combined position and intensity of the characteristic diffraction peaks rather than on isolated reflections.
The XRD analysis was used primarily for qualitative phase identification. A quantitative phase fraction or bulk reduction percentage was not calculated from the diffraction patterns because a full quantitative refinement was not performed. The results were instead compared across the two treatment temperatures and interpreted together with the SEM–EDS observations.
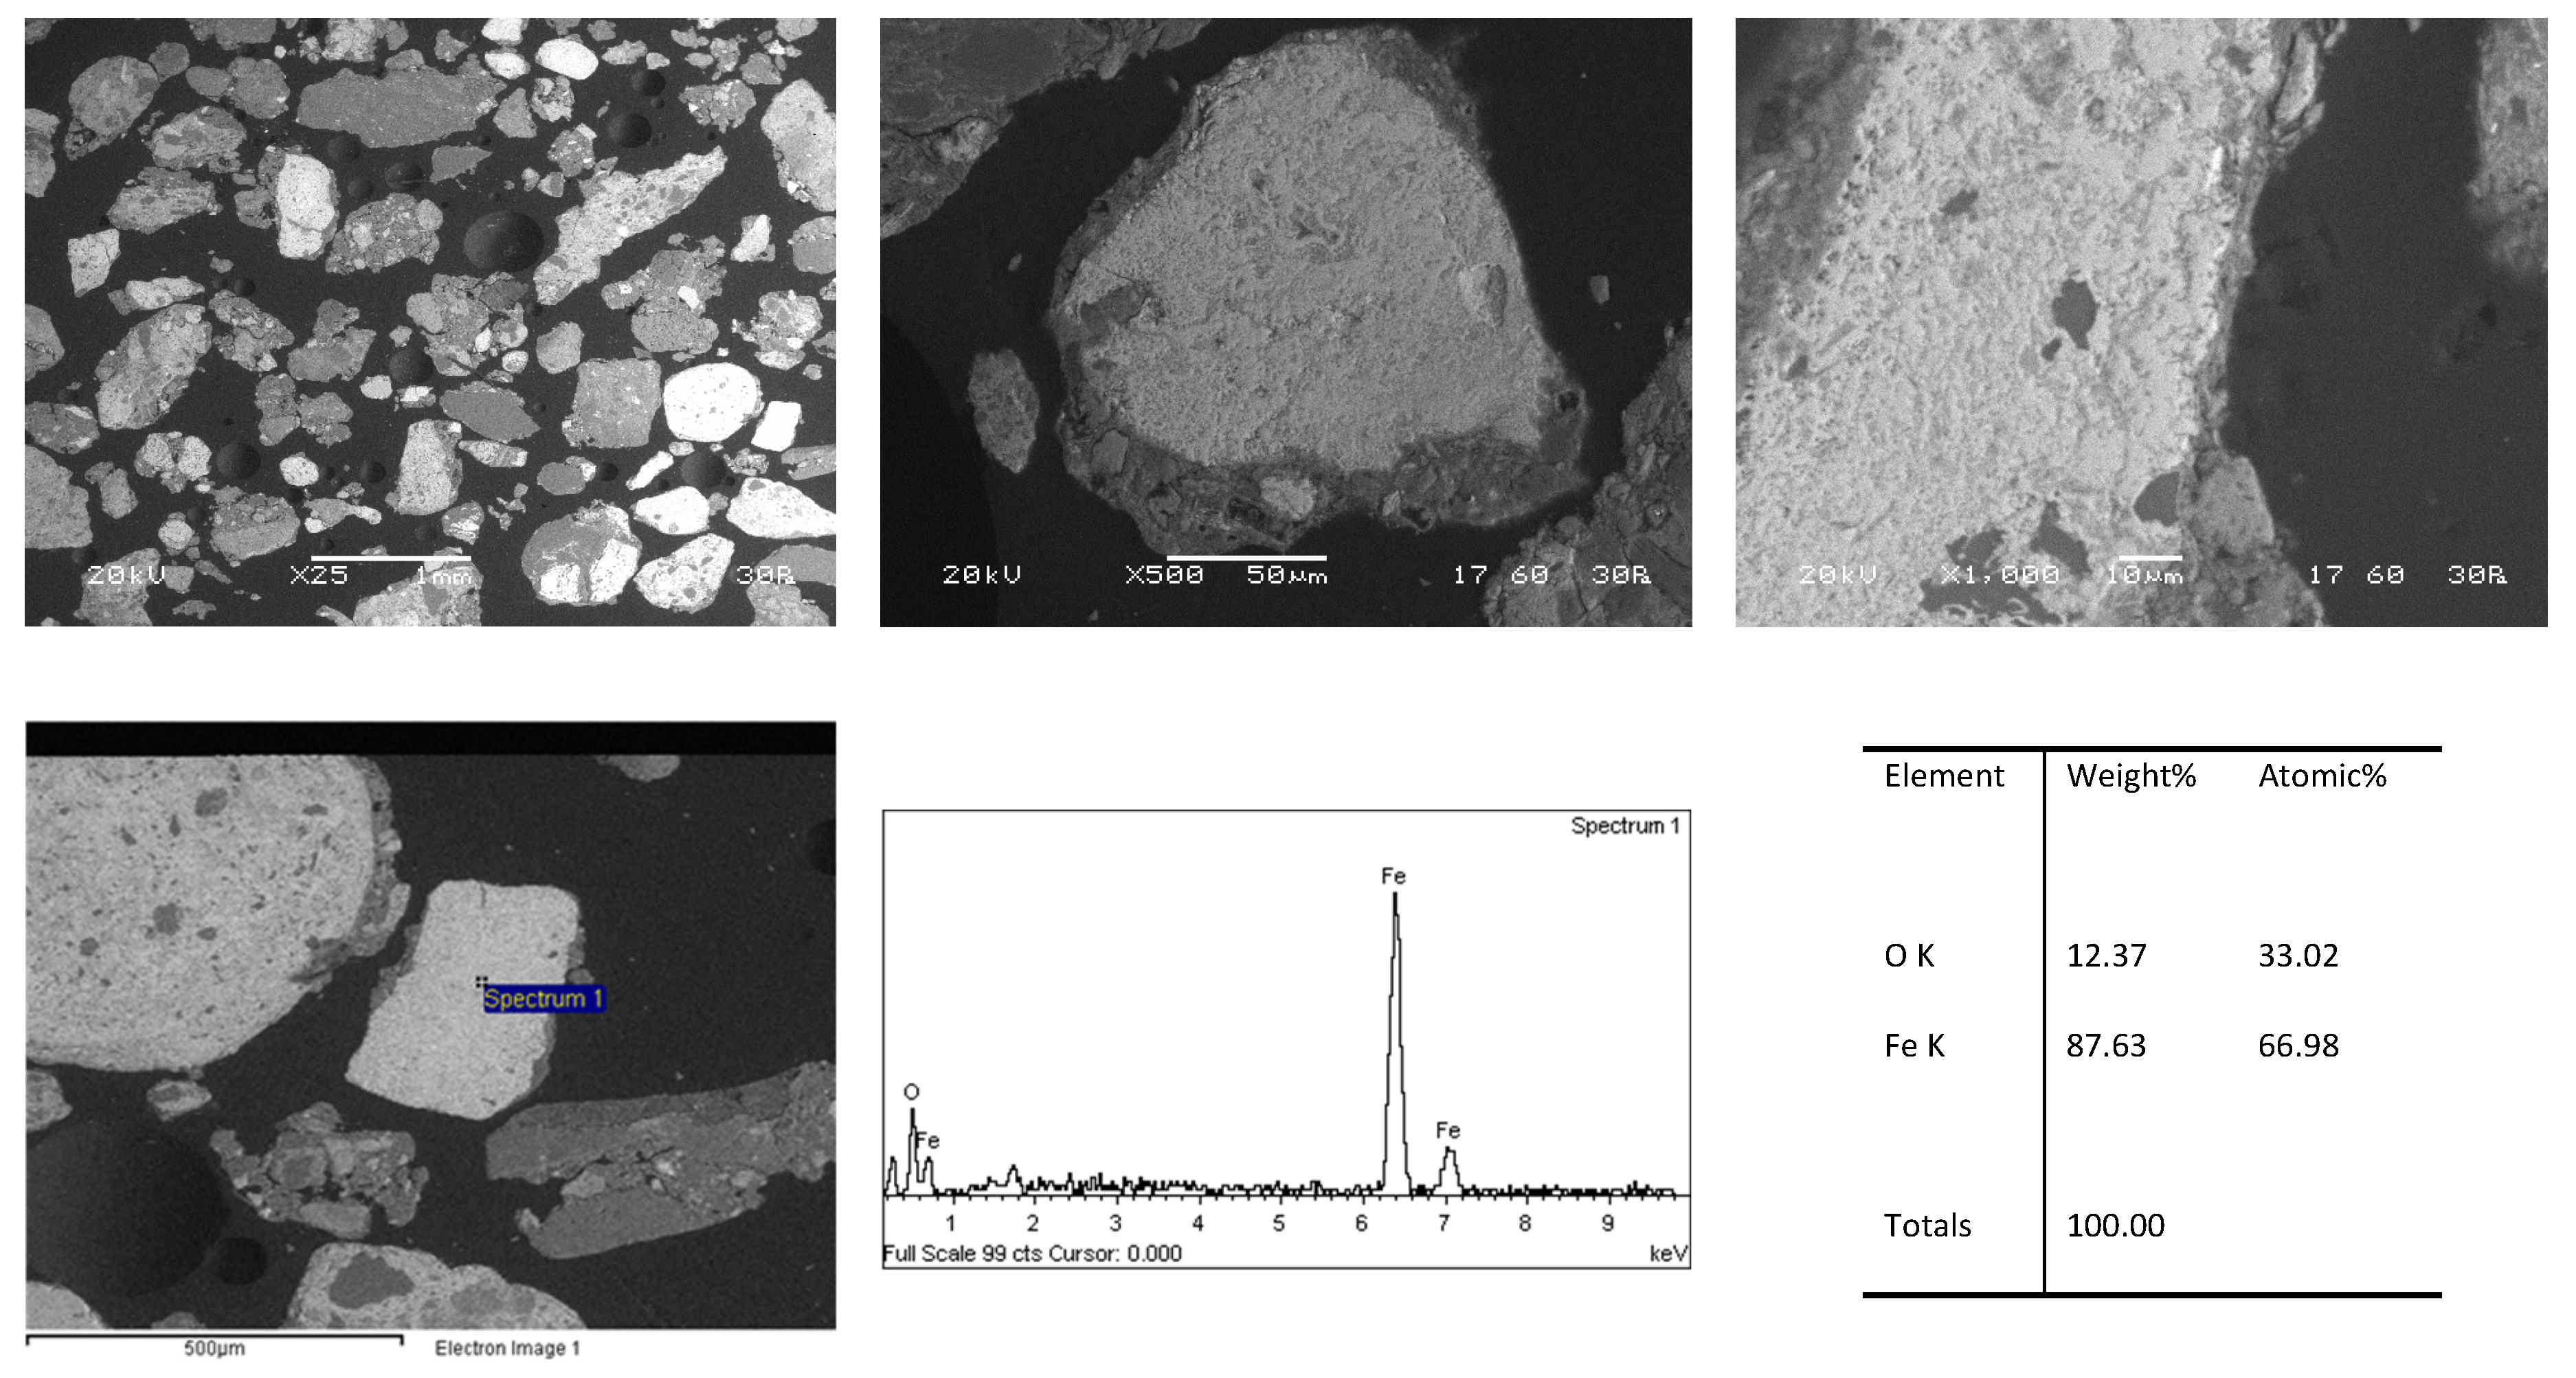
2.7.3. SEM–EDS Analysis
Scanning electron microscopy (SEM) coupled with energy-dispersive X-ray spectroscopy was used to examine the morphology and local elemental composition of the treated laterite samples. The analyses were performed on polished specimens obtained from the −1 mm fraction treated at 800 °C and 1000 °C. The samples were prepared by grinding and mechanical polishing with 6 μm and 1 μm diamond pastes before examination.
SEM images were assessed for changes in particle and grain morphology, the development of pores and microchannels, disruption of the oxide structure, the formation of dense or metallic regions, and evidence of heterogeneous or locally uniform reduction. The formation of internal pores and channels was considered in relation to the production and removal of water vapour during hydrogen reduction. These features may develop as oxygen is removed from the oxide lattice and gaseous H2O migrates out of the reacting grains. EDS point analyses and elemental maps were used to examine the local distribution of oxygen, iron, nickel, cobalt, chromium, magnesium, aluminium and silicon. The comparison focused on changes in oxygen intensity and on the development of Fe-, Ni- and Co-rich regions after treatment. The analyses were also used to examine whether the metallic elements were concentrated in isolated areas or distributed across the observed microstructure.
2.7.4. Comparative Analysis of Operating Variables
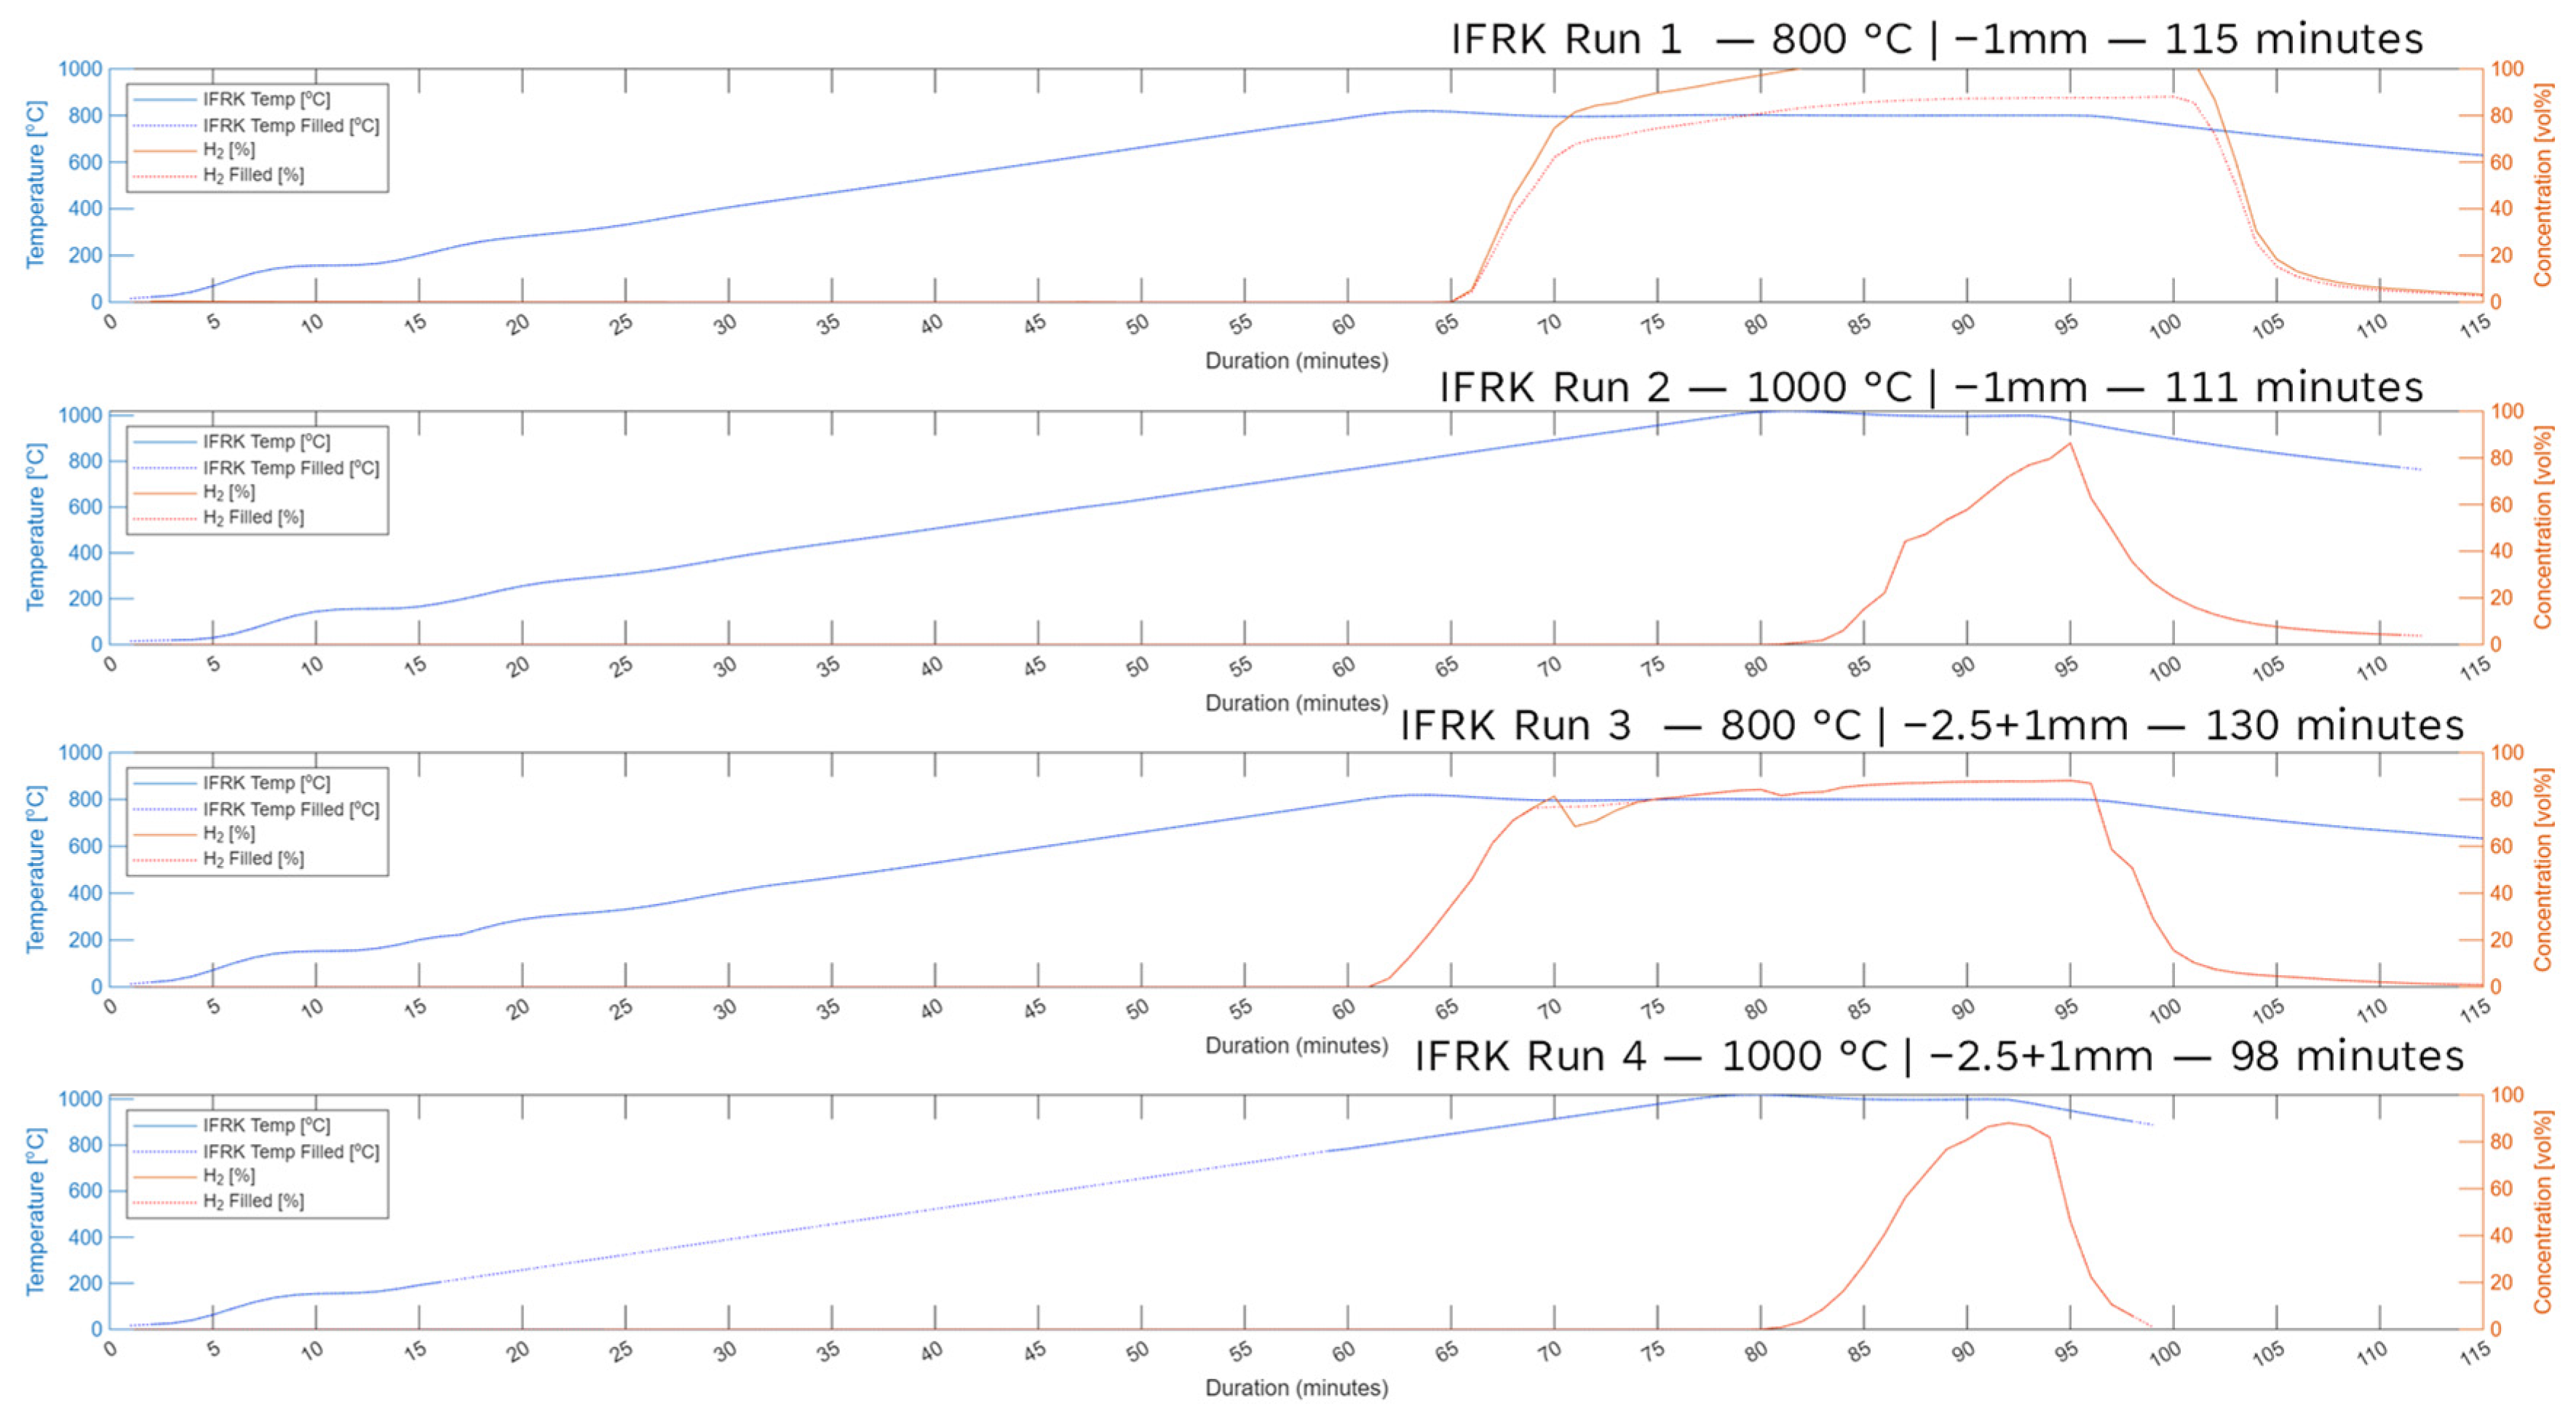
The reduction experiments were arranged to examine the combined influence of temperature and particle size. Two particle-size fractions were tested, including material finer than 1 mm and material between 1 and 2.5 mm. Each fraction was treated at 800 and 1000 °C. All experiments were conducted using the same kiln rotation speed (10 rpm) and the same nominal hydrogen flow rate (approximately 0.55 L/min), ensuring that temperature and particle size were the primary variables under investigation. The four runs therefore formed the comparison matrix shown in
Table 2.
The influence of temperature was examined by comparing runs performed with the same particle-size fraction. For the −1 mm fraction, the comparison was made between Run 1 and Run 2, while for the −2.5 + 1 mm fraction, the comparison was made between Run 3 and Run 4. The influence of particle size was examined by comparing runs performed at the same target temperature. At 800 °C, the comparison was made between Run 1 and Run 3, whereas at 1000 °C, the comparison was made between Run 2 and Run 4.
The comparative evaluation was based on the time required to reach the target temperature; the duration of hydrogen treatment; the H2 concentration profile after gas switching; hydrogen breakthrough and stabilisation behaviour; the calculated inlet, outlet and consumed hydrogen quantities; sample mass loss; XRD phase evolution, and SEM–EDS observations where available.
The duration of hydrogen treatment was not the same at the two temperatures. The tests at 800 °C were performed for 35 min, whereas the tests at 1000 °C were performed for 15 min. The temperature comparison therefore reflects the combined effect of temperature and treatment duration. The results were interpreted as experimental trends under the applied operating conditions rather than as an isolated kinetic effect of temperature.
2.7.5. Gas-Control Performance and H2 Indicators
The H2 concentration measured at the IFRK outlet was used to examine the transition from the nitrogen atmosphere to the hydrogen-reduction stage. The reduction experiments consisted of three main gas-related periods: heating of the sample under nitrogen, replacement of nitrogen by hydrogen at the target temperature, and termination of hydrogen feeding followed by nitrogen purging.
The beginning of hydrogen treatment was defined from the recorded gas-switching time and the subsequent increase in outlet H2 concentration. The initial increase in measured H2 represents the combined effects of gas transport, nitrogen displacement, filling of the kiln and connecting lines, analyser response and hydrogen consumption by the reacting material. The concentration increase was therefore not interpreted independently as a direct reduction rate.
The synchronised temperature and H
2 profiles were used to identify the time at which hydrogen was introduced, the first detectable H
2 response at the analyser, the period of rapid H
2 increase, the time required to reach selected H
2 concentration levels, the period of H
2 stabilisation, and the final decrease associated with the termination of hydrogen supply and nitrogen purging. Hydrogen breakthrough time was defined as the interval between the recorded introduction of hydrogen and the first sustained increase in the outlet H
2 concentration. The times required for the signal to reach 50%, 80% and 90% of its stable value were also determined where the recorded profile permitted this calculation. The local H
2 concentration slope (S
H2) was calculated as
where ΔC
H2 is the change in H
2 concentration, and Δt is the corresponding change in time. The result was expressed in concentration percentage points per minute. Positive values represent an increase in the measured outlet H
2 concentration, whereas negative values represent a decrease. These slopes describe changes in the measured gas signal and were not treated as direct molar reaction rates.
The total volume of hydrogen supplied during each experiment was calculated from the measured or nominal inlet flow and the hydrogen-treatment duration:
where V
H2,in is the total hydrogen volume supplied (L), Q
H2,in is the average hydrogen inlet-flow rate (L/min) and t
H2 is the hydrogen-treatment time (min).
The corresponding hydrogen mass was calculated using the hydrogen density adopted in the experimental analysis:
where m
H2,in is the hydrogen mass supplied (g) and ρ
H2 is the hydrogen density (g/L). A density of ρ
H2 = 0.08988 g/L was used for the conversion of hydrogen volume to mass.
The outlet hydrogen quantity was estimated from the measured H
2 concentration profile and the gas-flow information used in the experimental analysis. The outlet hydrogen volume, V
H2,out, was calculated by integrating the hydrogen fraction in the measured gas stream over the selected hydrogen-treatment period. The corresponding hydrogen mass, m
H2,out, was calculated as
where m
H2,out is the outlet hydrogen mass (g), V
H2,out is the outlet hydrogen volume (L) and ρ
H2 is the hydrogen density (g/L). The outlet quantity represents the hydrogen detected in the measured gas stream during the selected period.
The apparent hydrogen consumption (V
H2,cons) was calculated as the difference between the inlet and outlet quantities:
Similarly, the apparent hydrogen mass consumption was calculated as
The fraction of the supplied hydrogen classified as consumed was calculated as
where η
H2 is the apparent hydrogen consumption (%).
This indicator was used to compare the four experimental runs. It was described as apparent hydrogen consumption because the calculation does not separately quantify all gas accumulation and displacement effects occurring in the kiln, connecting lines and analyser circuit during the transition from nitrogen to hydrogen.
During data processing, isolated missing data points resulting from brief interruptions in signal acquisition were treated using interpolation, only when necessary to preserve the continuity of the recorded time series for graphical presentation and trend interpretation. These interpolated points were not used to alter or influence the quantitative evaluation of the experimental results, and the main conclusions of the study remained unchanged when considering only the directly measured data. No experimental observations were intentionally excluded from the analysis. Accordingly, a clear distinction was maintained between genuinely missing measurements caused by temporary acquisition interruptions and censored data, the latter of which were not present in this experimental campaign.
4. Discussion
The present work demonstrates that the improved performance achieved during both magnesite calcination and hydrogen-assisted laterite reduction originates from the combined operation of an airtight reactor and an electrified IFRK rather than from either feature individually. Conventional rotary kilns rely on combustion gases to provide both heat and the reaction atmosphere, making thermal control and gas composition intrinsically coupled. In contrast, the proposed reactor physically separates heat generation from the reacting material while maintaining a sealed reaction chamber, allowing temperature and atmosphere to be controlled independently throughout the process. This operating principle creates a stable thermochemical environment in which heat transfer is more uniform, air infiltration is eliminated and the composition of the process gas remains essentially constant. Such conditions become increasingly important as the metallurgical industry shifts towards hydrogen-based processing, where even small changes in oxygen partial pressure or hydrogen concentration can influence reaction pathways, gas utilisation and product quality.
The benefits of this integrated approach are evident in both investigated processes. During the calcination of magnesite, the combination of uniform indirect heating, continuous mixing, and a controlled atmosphere sped up the breakdown of carbonates, leading to a conversion rate of over 98% in much shorter treatment times than what is usually needed in industrial rotary kilns. The shorter residence time is not solely a consequence of higher operating temperature but also reflects improved reaction conditions inside the rotary kiln. Continuous removal of the evolved CO2 limits local equilibrium effects that can retard decomposition, while the homogeneous thermal field ensures that particles experience similar heating histories throughout the rotating bed. Together, these factors enhance reaction kinetics, reduce unnecessary heat losses associated with prolonged operation and increase reactor productivity without compromising product quality. The results therefore suggest that improving the reaction environment can be as important as increasing the supplied thermal energy when seeking higher calcination efficiency.
The same reactor characteristics proved even more beneficial during hydrogen-assisted laterite reduction. Unlike conventional systems, where hydrogen may be diluted by combustion products or consumed through air infiltration, the airtight configuration preserves a stable reducing atmosphere throughout the treatment. Consequently, hydrogen is primarily used for oxide reduction, improving both process efficiency and hydrogen utilisation. This effect becomes particularly evident when comparing the two operating temperatures. Increasing the reduction temperature from 800 °C to 1000 °C reduced the required hydrogen treatment time from 35 min to only 15 min, demonstrating the strong acceleration of reduction kinetics once favourable thermodynamic conditions are combined with efficient heat transfer and a stable hydrogen atmosphere. This short treatment duration compares favourably with earlier hydrogen-reduction studies of lateritic ores, in which treatment periods exceeding three hours were reported under additive-assisted conditions [
17]. Such a substantial reduction in residence time directly increases throughput while simultaneously lowering the total hydrogen requirement and the electrical energy associated with prolonged furnace operation. Since hydrogen cost remains one of the principal challenges for industrial implementation of low-carbon metallurgy, reducing both hydrogen consumption and treatment time represents a significant technological advantage.
Beyond the experimental results themselves, the study highlights the importance of combining electrification with atmosphere isolation as an innovative decarbonised technology for high-temperature metallurgical reactors. Electrification alone cannot fully exploit the benefits of hydrogen if the reaction atmosphere remains poorly controlled, while airtightness alone cannot deliver the rapid and responsive thermal management offered by electrical heating. The synergy between these two concepts enables precise control of both thermal and chemical conditions, creating favourable conditions for faster reaction kinetics, improved gas efficiency and greater process flexibility. Furthermore, because the exhaust gas is not diluted by combustion products, the reactor provides favourable conditions for future hydrogen recovery and recirculation, offering an additional pathway for reducing operating costs and improving overall process sustainability. Although demonstrated at laboratory scale, these results indicate that airtight electrified rotary kilns could provide an effective platform for next-generation calcination and hydrogen-reduction processes, supporting the transition towards highly efficient, low-carbon and renewable electricity-driven metallurgical production.
5. Conclusions
This study demonstrated the laboratory-scale feasibility of an airtight electrified indirect-fired rotary kiln for two distinct thermochemical applications: magnesite calcination, and hydrogen reduction of lateritic ore. The reactor maintained controlled thermal and atmospheric conditions while enabling synchronised monitoring of temperature, gas composition and flow-related variables.
Magnesite calcination was successfully followed through the evolution and depletion of CO2. Higher treatment temperatures shortened the overall process duration and produced a sharper measurable gas response. TG–DTA confirmed extensive carbonate decomposition, while the mass-balance and particle-size results showed that calcination was accompanied by substantial fragmentation and some physical product loss.
During laterite reduction, the reactor achieved the required sequence of nitrogen heating, hydrogen treatment and final purging. The gas-phase results indicated substantial apparent hydrogen interaction, while XRD and SEM–EDS confirmed progressive transformation of iron- and nickel-bearing phases. Reduction remained partial at 800 °C and advanced further at 1000 °C, where metallic Fe- and Ni-rich regions became more evident, although residual oxides and silicate-bound phases persisted. The results highlight the potential benefits of hydrogen as a reducing agent, including the possibility of accelerating reduction kinetics, improving metal liberation and selectivity, and reducing direct carbon-related emissions compared with conventional reductants. These characteristics make hydrogen a promising pathway for optimising future laterite processing routes, particularly when coupled with electrified heating systems.
The main contribution of the work is the integration of indirect electrical heating, sealed rotary operation, controlled gas atmospheres and synchronised process monitoring within a common experimental platform. The results also confirm that online gas measurements must be combined with solid characterisation to assess reaction progress reliably. Further development should focus on complete outlet-gas measurement, direct monitoring of water vapour, improved bed-temperature characterisation and replicated experimental campaigns. These improvements are required to support quantitative process modelling, optimisation, scale-up and future digital twin applications.