


Abstract
The TRM (TONNEAU1 Recruiting Motif) gene family plays a crucial role in multiple biological processes, including microtubule organization, cell division regulation, fruit morphogenesis, stress adaptation, and growth and development. To delve deeper into the potential functions of BnaTRMs in Brassica napus, this study employed bioinformatics methods to systematically identify and analyze the TRM family genes in Brassica napus (Westar). Using the model plant Arabidopsis thaliana as a reference and based on six conserved motifs, 100 TRM members were first identified in Brassica napus. These genes are widely distributed across 19 chromosomes, and most exhibit nuclear localization characteristics. Through gene collinearity analysis among Brassica napus, Arabidopsis thaliana, Glycine max, Oryza sativa, and Zea mays, we speculate that Brassica napus and Glycine max may share a similar evolutionary history. Analysis of cis-acting elements in the 2000 bp upstream region of TRM gene promoters revealed numerous elements related to abiotic stress response and hormone regulation. Furthermore, qRT-PCR data supported these findings, indicating that multiple TRM genes actively participate in the growth and development process and abiotic stress tolerance of Brassica napus. In summary, BnaTRMs exhibit significant functions in stress adaptation, growth, and development. This study not only enhances our understanding of the functions of the TRM gene family but also provides new perspectives and strategies for further exploring their regulatory mechanisms and potential applications.
1. Introduction
Brassica napus L. is one of the vital oilseed crops in China [1], the allotetraploid rapeseed (AnAnCnCn, 2n = 4x = 38) originates from spontaneous hybridization of its diploid ancestors Brassica rapa (ArAr, 2n = 2x = 20) and Brassica oleracea (CoCo, 2n = 2x = 18) [2,3,4]. In agricultural ecology, the root system of Brassica napus L. plays a crucial role in improving soil structure, enhancing soil fertility, and preventing water and soil loss. Additionally, planting Brassica napus L. can effectively suppress weed growth, thereby reducing pesticide usage and benefiting ecological conservation. The development of the Brassica napus L. industry is of great significance in ensuring national food security, promoting economic growth, and maintaining ecological balance.
The TRM gene family plays crucial roles in plant growth and development. In Arabidopsis thaliana, 34 TRM proteins have been identified, half of which are microtubule-associated proteins involved in microtubule organization formation and maintenance [5,6]. Charge analysis of TRM primary sequences reveals that approximately half of TRM family members contain large basic regions, suggesting these proteins likely participate in microtubule binding [7,8]. Microtubules serve as essential structural elements within the plant cytoskeleton, fulfilling critical functions in cellular morphology maintenance and adaptive responses during growth, developmental processes, and environmental fluctuations. Additionally, they participate in fundamental cellular activities, including mitotic division, organelle trafficking, pathogen defense mechanisms, and stress adaptation [9,10]. TONNEAU1 (TON1) protein represents a highly conserved acidic protein in land plants [11]. TONNEAU1 Recruiting Motif (TRM) proteins derive their nomenclature from their functional association with TON1, a plant-specific protein homologous to the human centrosomal FOP protein that is critically required for proper microtubule array formation in Arabidopsis thaliana [5]. The TON1 proteins serve as a crucial regulator of leaf and silique morphology in Arabidopsis thaliana [12], grain shape in Oryza sativa L. [13,14], as well as fruit morphology in Solanum lycopersicum and Cucumis sativus L. [15]. The TRM proteins interact with TON1 and protein phosphatase 2A (PP2A) through their M2 and M3 domains, respectively, forming the TON1-TRM-PP2A (TTP) protein complex. This complex specifically localizes to microtubules (MTs), where it regulates microtubule organization and preprophase band (PPB) formation, thereby modulating cell division and growth processes that ultimately determine the size and morphology of plant organs [6,16,17,18,19,20].
In Arabidopsis thaliana, the Attrm5 mutant exhibits retarded leaf growth, delayed flowering, and reduced root length [21]. AtTRM61 possesses conserved functional domains, including a characteristic motif for S-adenosyl-L-methionine (AdoMet) cofactor binding, which plays a crucial role in regulating embryonic arrest and seed abortion processes [22]. The research demonstrates that AtTRM1 and AtTRM2 regulate leaf morphology by actively promoting longitudinal polar cell elongation [5]. These two genes play crucial roles in determining final leaf shape and size through modulating cell elongation patterns. This regulatory mechanism reveals the precise control function of TRM family genes in plant organ morphogenesis, providing important clues for understanding the molecular mechanisms of plant organ development. Recent studies show that AtTRM21 positively regulates flavonoid biosynthesis at the translational level in Arabidopsis thaliana. Loss-of-function mutation in TRM21 leads to root hair growth defects and delayed plant growth, accompanied by significant alterations in secondary metabolites, particularly a marked reduction in flavonoid content. This discovery not only reveals a novel metabolic regulatory function of TRM21 but also provides new molecular targets for elucidating the regulatory network of flavonoid biosynthesis [23]. In rice, TRM homologous gene OsGW7/GL7/SLG7 ultimately determines grain size and quality traits by regulating cell length and width [24,25]. In Solanum lycopersicum, TRM protein family members interact with the microtubule organizing center OFP to regulate fruit morphology by controlling cell division orientation and cell expansion. OVATE and SlOFP20 interact with SlTRM5 and ten other TRM family members. The TRM-OFP protein complexes exhibit dynamic subcellular localization in the cytoplasm and microtubules, dependent on their co-expression patterns, suggesting that their spatial distribution may contribute to morphological regulation [26,27,28,29]. The research demonstrates that this TRM-OFP interaction network precisely coordinates cellular behaviors during fruit development, ultimately determining final fruit shape. In Cucumis sativus L., some CsTRM genes are induced or suppressed at different time points under stress treatments. Particularly, CsTRM21 shows significant expression variations between long and short fruited cultivars, as well as under abiotic stresses (salt and heat) and biotic stresses (powdery mildew and gray mold), suggesting its dual role in determining fruit shape and stress resistance [30].
As a crucial oilseed crop, rapeseed (Brassica napus L.) is particularly susceptible to abiotic stresses, including chilling stress, drought stress, and salt stress, which significantly impair its growth dynamics and developmental processes and ultimately compromise both yield potential and oil quality traits [31]. Under abiotic stress conditions, the dynamic reorganization of the microtubule cytoskeleton plays a critical role in maintaining cellular homeostasis [32]. These three abiotic stresses can significantly disrupt the structure and function of microtubules in plant cells, thereby interfering with critical physiological processes, including cell division, cell expansion, cell morphogenesis, and intracellular transport. Under cold stress conditions, low temperatures destabilize the polymerization equilibrium of microtubule subunits, leading to aberrant cell division patterns that ultimately manifest as phenotypic abnormalities such as leaf wrinkling and organ deformation [33]. Under drought stress conditions, microtubules mediate stomatal closure, consequently reducing transpiration rates and compromising cellular osmoregulatory capacity (microtubule-associated protein AtMPB2C plays a role in the organization of cortical microtubules, stomata patterning, and tobamovirus infectivity). Under high-salinity stress conditions, cortical microtubules in plant cells undergo depolymerization, disrupting microtubule dynamic equilibrium and consequently impairing tissue development and organ morphogenesis [34]. Notably, these stresses often exert synergistic effects on the microtubule system. For instance, the combined action of salinity–alkalinity and drought stress significantly exacerbates microtubule network disorganization, while low-temperature stress further amplifies these disruptive effects. Members of the TRM gene family serve as crucial regulators of plant growth and stress responses, playing pivotal roles in plant morphogenesis and stress adaptation through their involvement in microtubule organization and cell division regulation. The evolutionary conservation of this gene family across diverse plant species makes it an important target for crop genetic improvement. However, to date, there have been no reported studies investigating the participation of BnaTRM family genes in abiotic stress responses in rapeseed.
In this study, we systematically identified 100 BnaTRM genes with high homology to Arabidopsis thaliana in the Brassica napus Westar cultivar using bioinformatics approaches. Comprehensive analyses were conducted to characterize their structural features and chromosomal distributions, and a multi-species phylogenetic tree was constructed, including Arabidopsis thaliana, Glycine max, Oryza sativa, and Zea mays. Furthermore, quantitative expression profiling revealed stress-specific expression patterns among BnaTRM family members, with several key genes exhibiting distinctive regulatory profiles that suggest functional specialization in response to multiple stresses. Our findings not only fill a critical knowledge gap in rapeseed TRM gene family research but also establish a foundation for investigating the potential roles of TRM in stress tolerance mechanisms in Brassica napus.
2. Results
2.1. One Hundred TON1 Recruiting Motif Family Members in Westar Were Mainly Divided into Eight Subfamilies
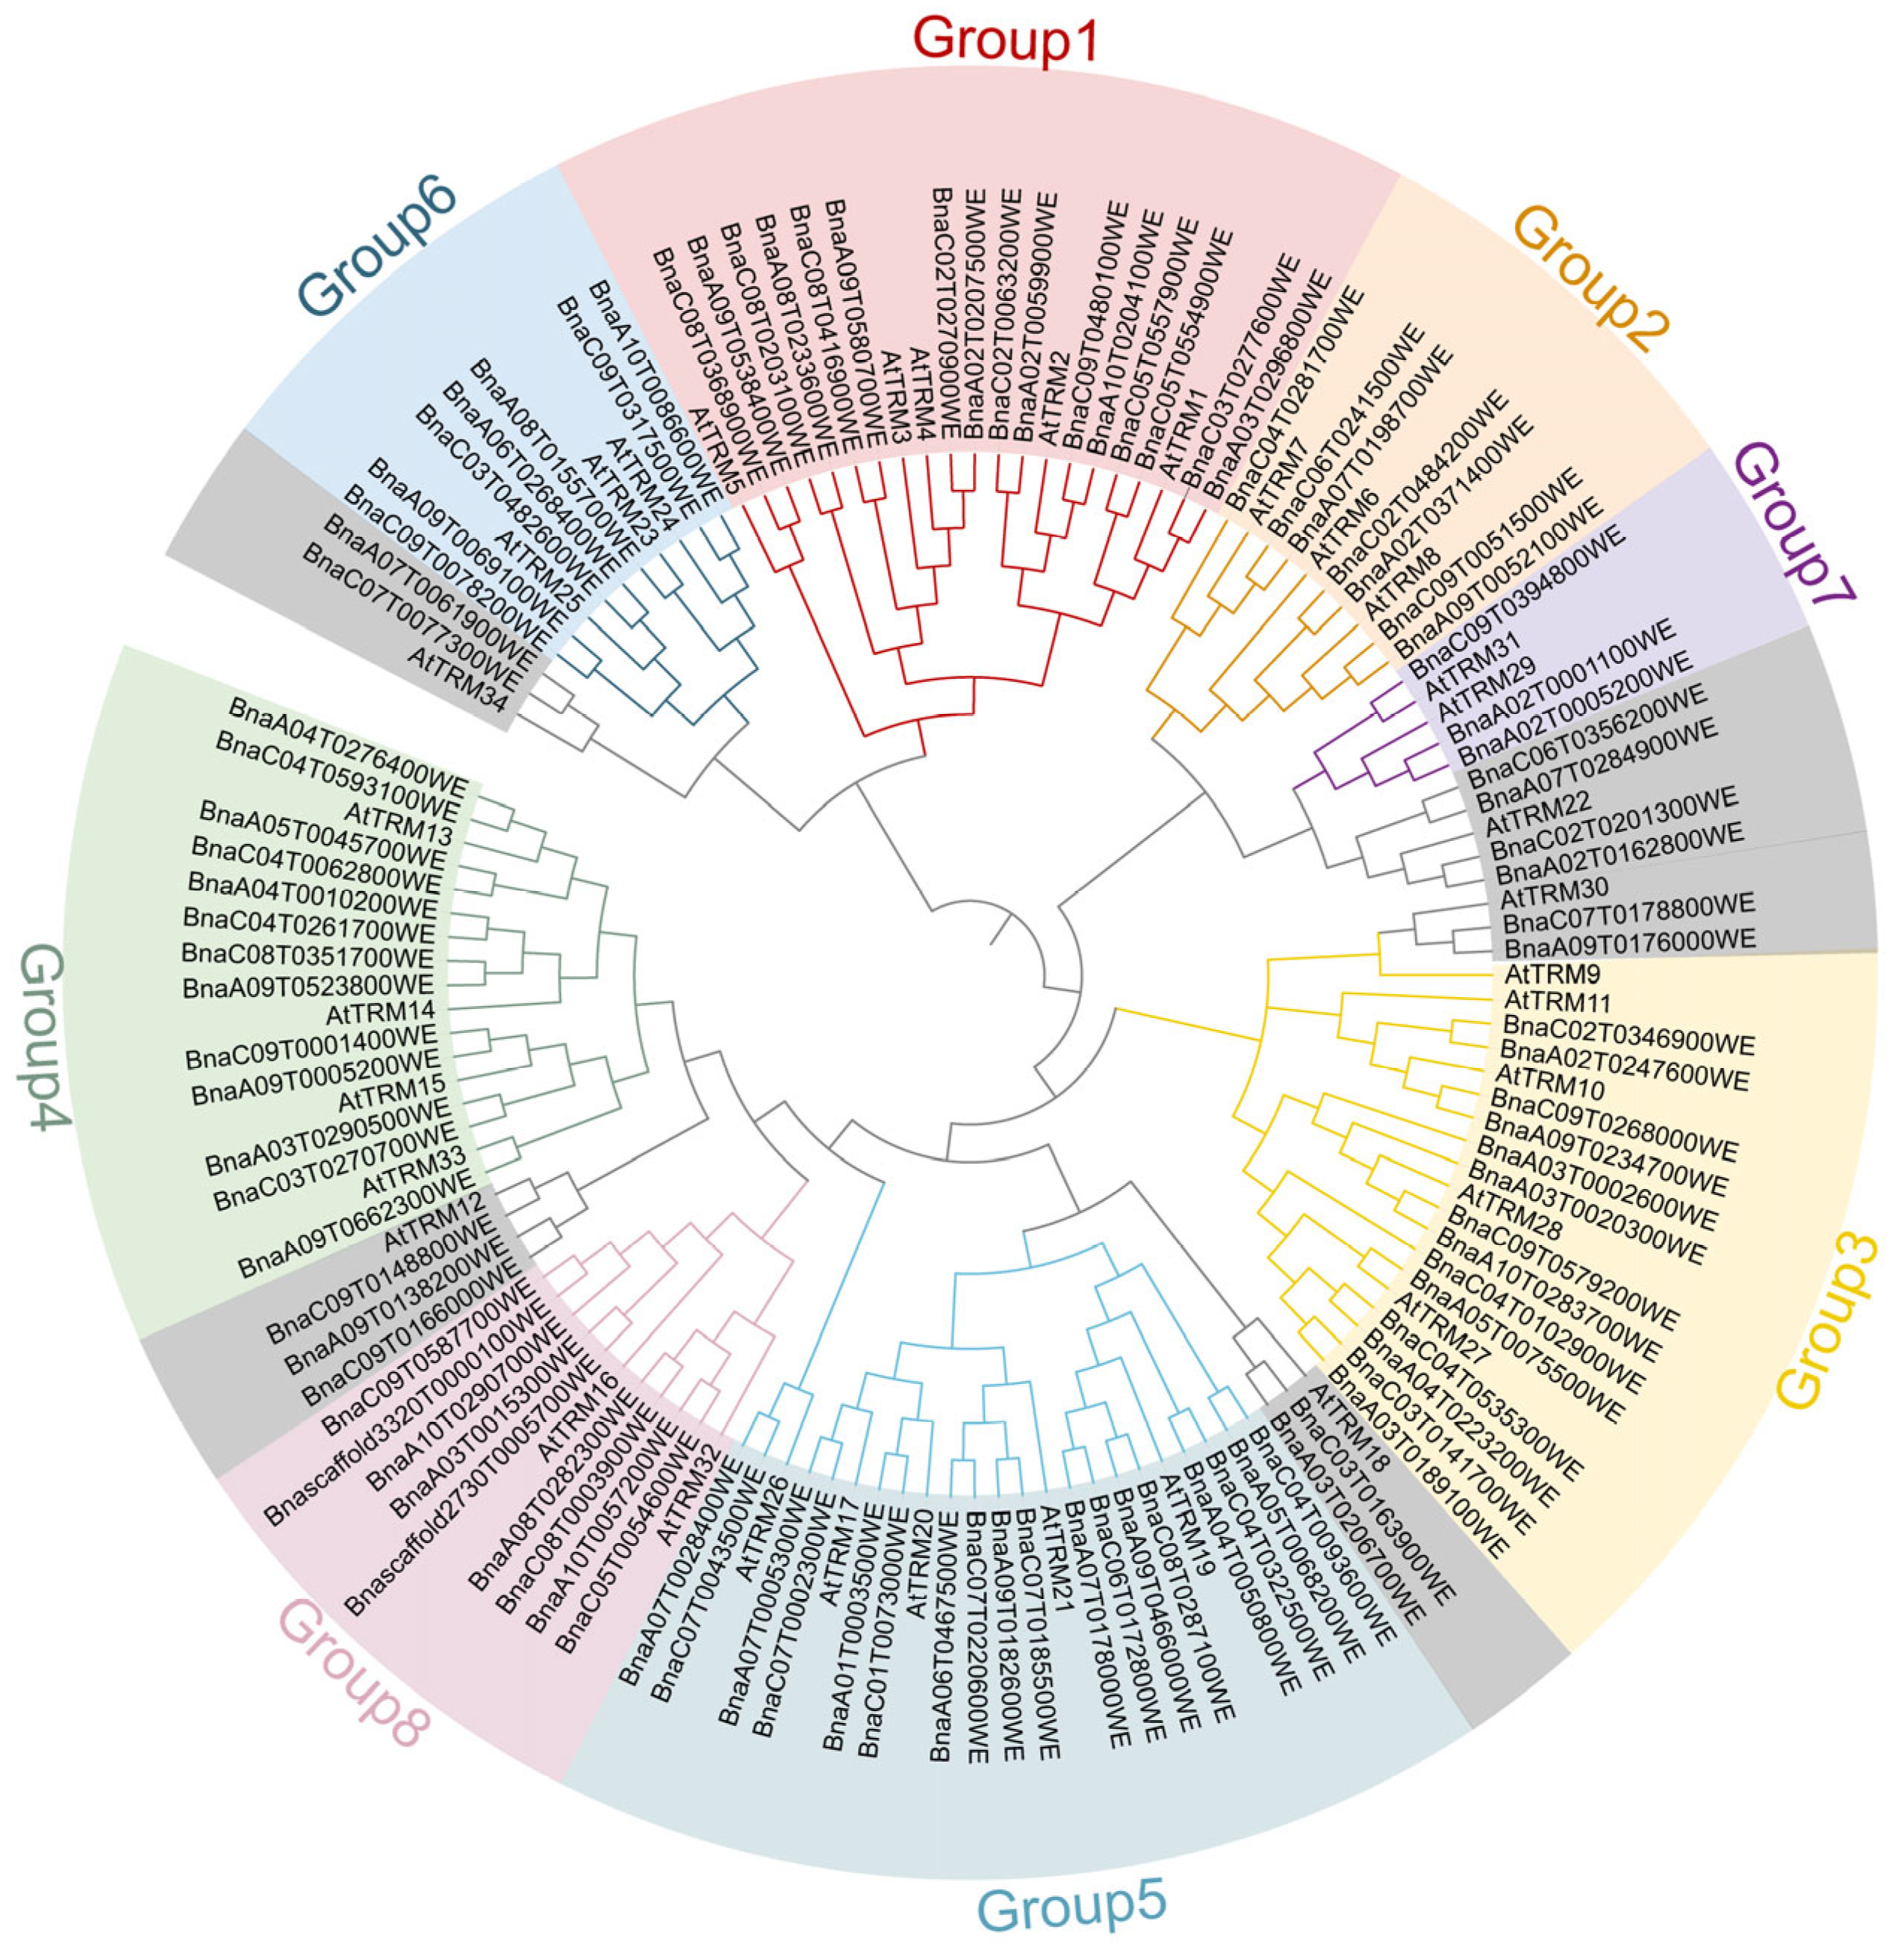
The TON1 Recruiting Motif (TRM) has been identified in Arabidopsis thaliana, a cruciferous plant. Consequently, using the amino acid sequences of AtTRMs as the source, a Blast search was conducted against the Brassica napus genome database, yielding 128 genes with E-values less than 1.0 × 10⁻⁵. Using the MEME v5.5.7 database to analyze their motifs, those with the conserved motifs were designated members of the TRM gene family. Thus, 100 TRM family members were identified (Table 1). To further understand the evolutionary relationships among the TRM family members in Brassica napus, comparative sequence alignments of TRM genes were conducted for Brassica napus and Arabidopsis thaliana, with subsequent generation of an unrooted phylogenetic tree utilizing MEGA 11.0.13 software. As shown in Figure 1, the rapeseed TRM family is divided into eight subfamilies and some independent branches. The largest clade, Group 3, comprises 18 BnaTRM members and five AtTRM proteins. In contrast, the smallest clade, Group 7, contains three BnaTRM and two AtTRM proteins (Figure 1).
Table 1.
Information regarding the BnaTRMs.
Figure 1.
Phylogenetic analysis of the TRM family genes of Brassica napus and Arabidopsis thaliana. The protein sequence alignments and construction of the phylogenetic tree were performed using MEGA 11.0.13 and the neighbor-joining method with 1000 bootstrap replicates. The different colors represent eight subfamilies of the TRM gene family; branches indicate different evolutionary clades. Gray represents independent branches.
The remarkable expansion of the TRM gene family in Brassica napus (containing 100 members) may be intrinsically linked to whole-genome duplication events during polyploidization. The retention of 652 homologous gene pairs between the A/C subgenomes likely facilitated functional module differentiation [35,36] (e.g., stress response-related subfamilies) and redundancy buffering mechanisms, thereby synergistically reinforcing adaptive evolutionary advantages in response to complex environmental stresses.
2.2. Gene Motif and Structure of the BnaTRMs
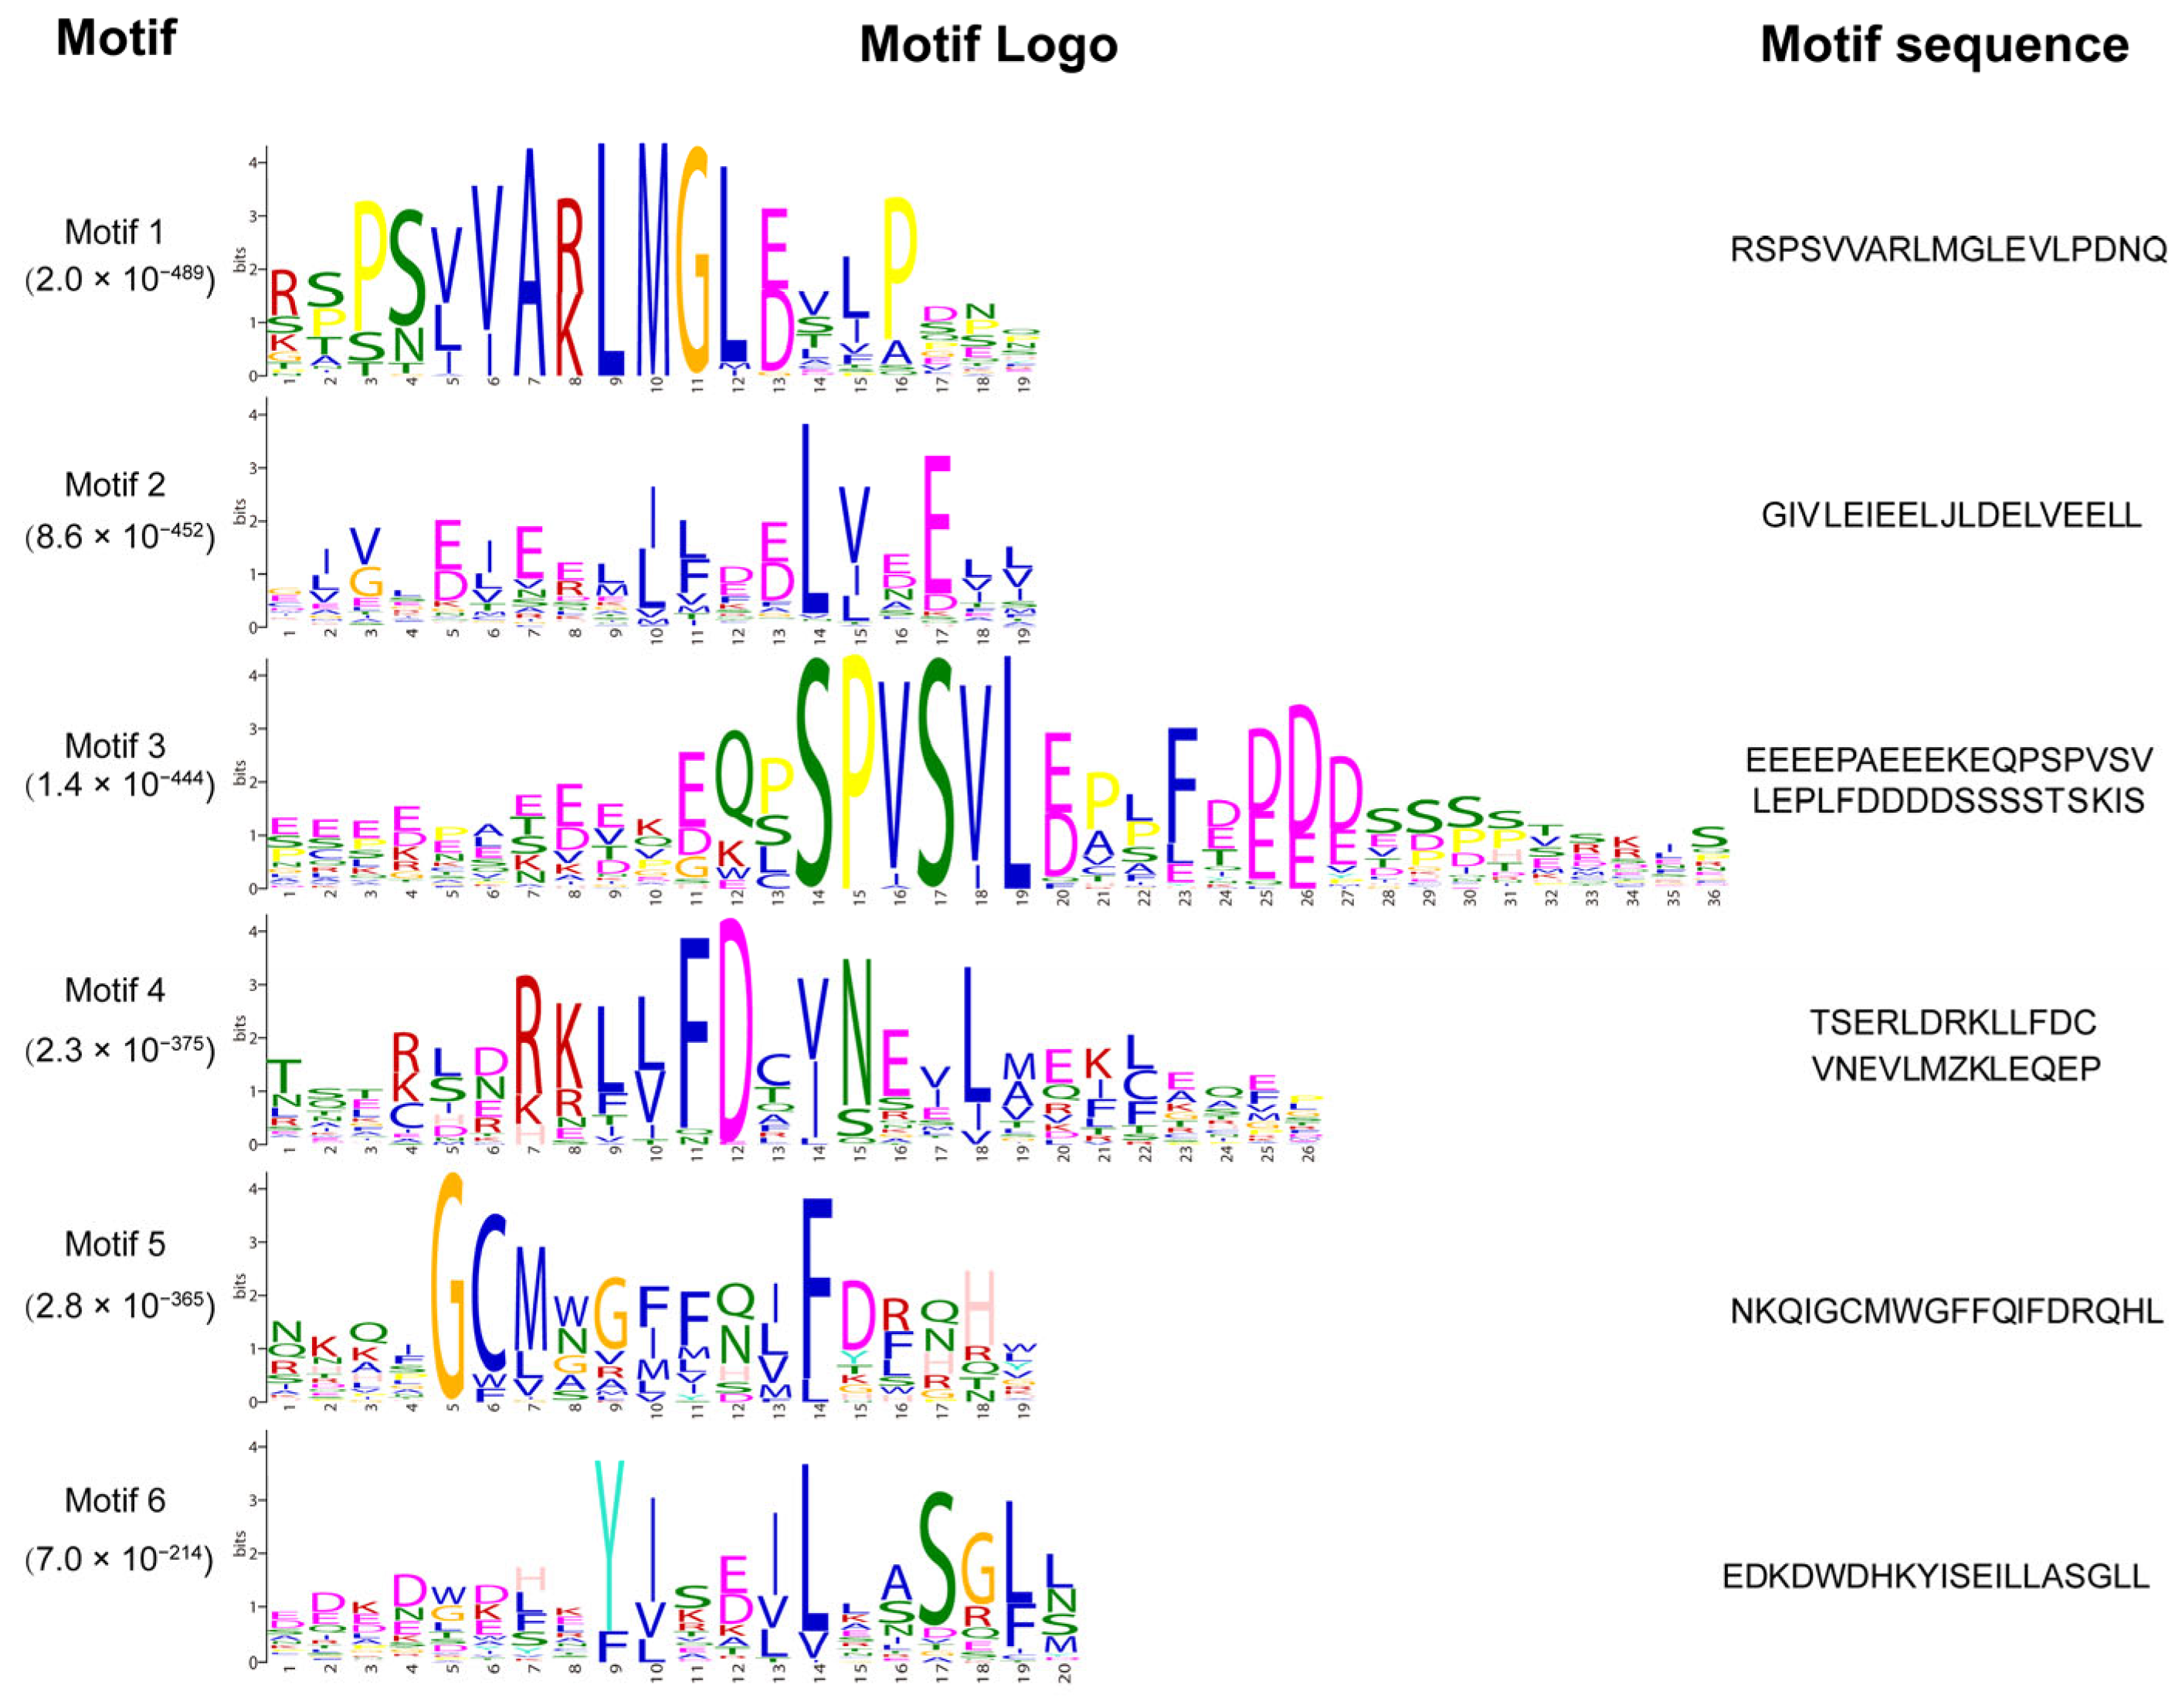
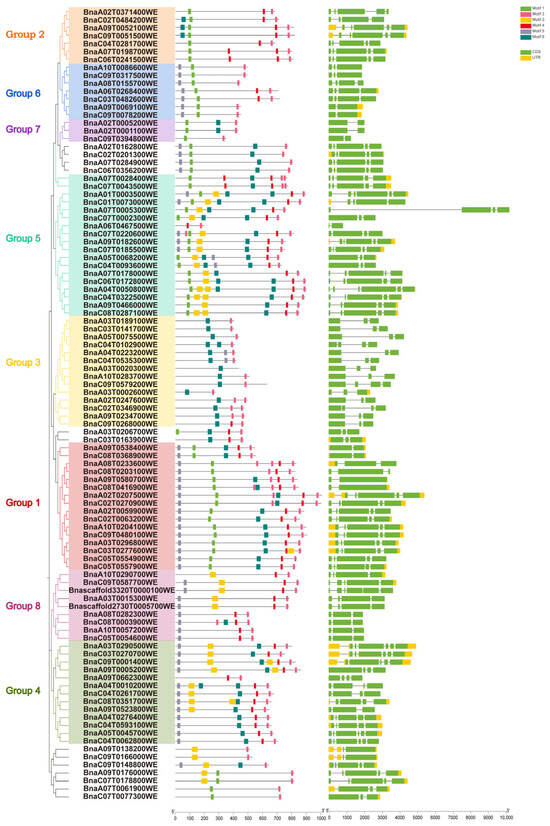
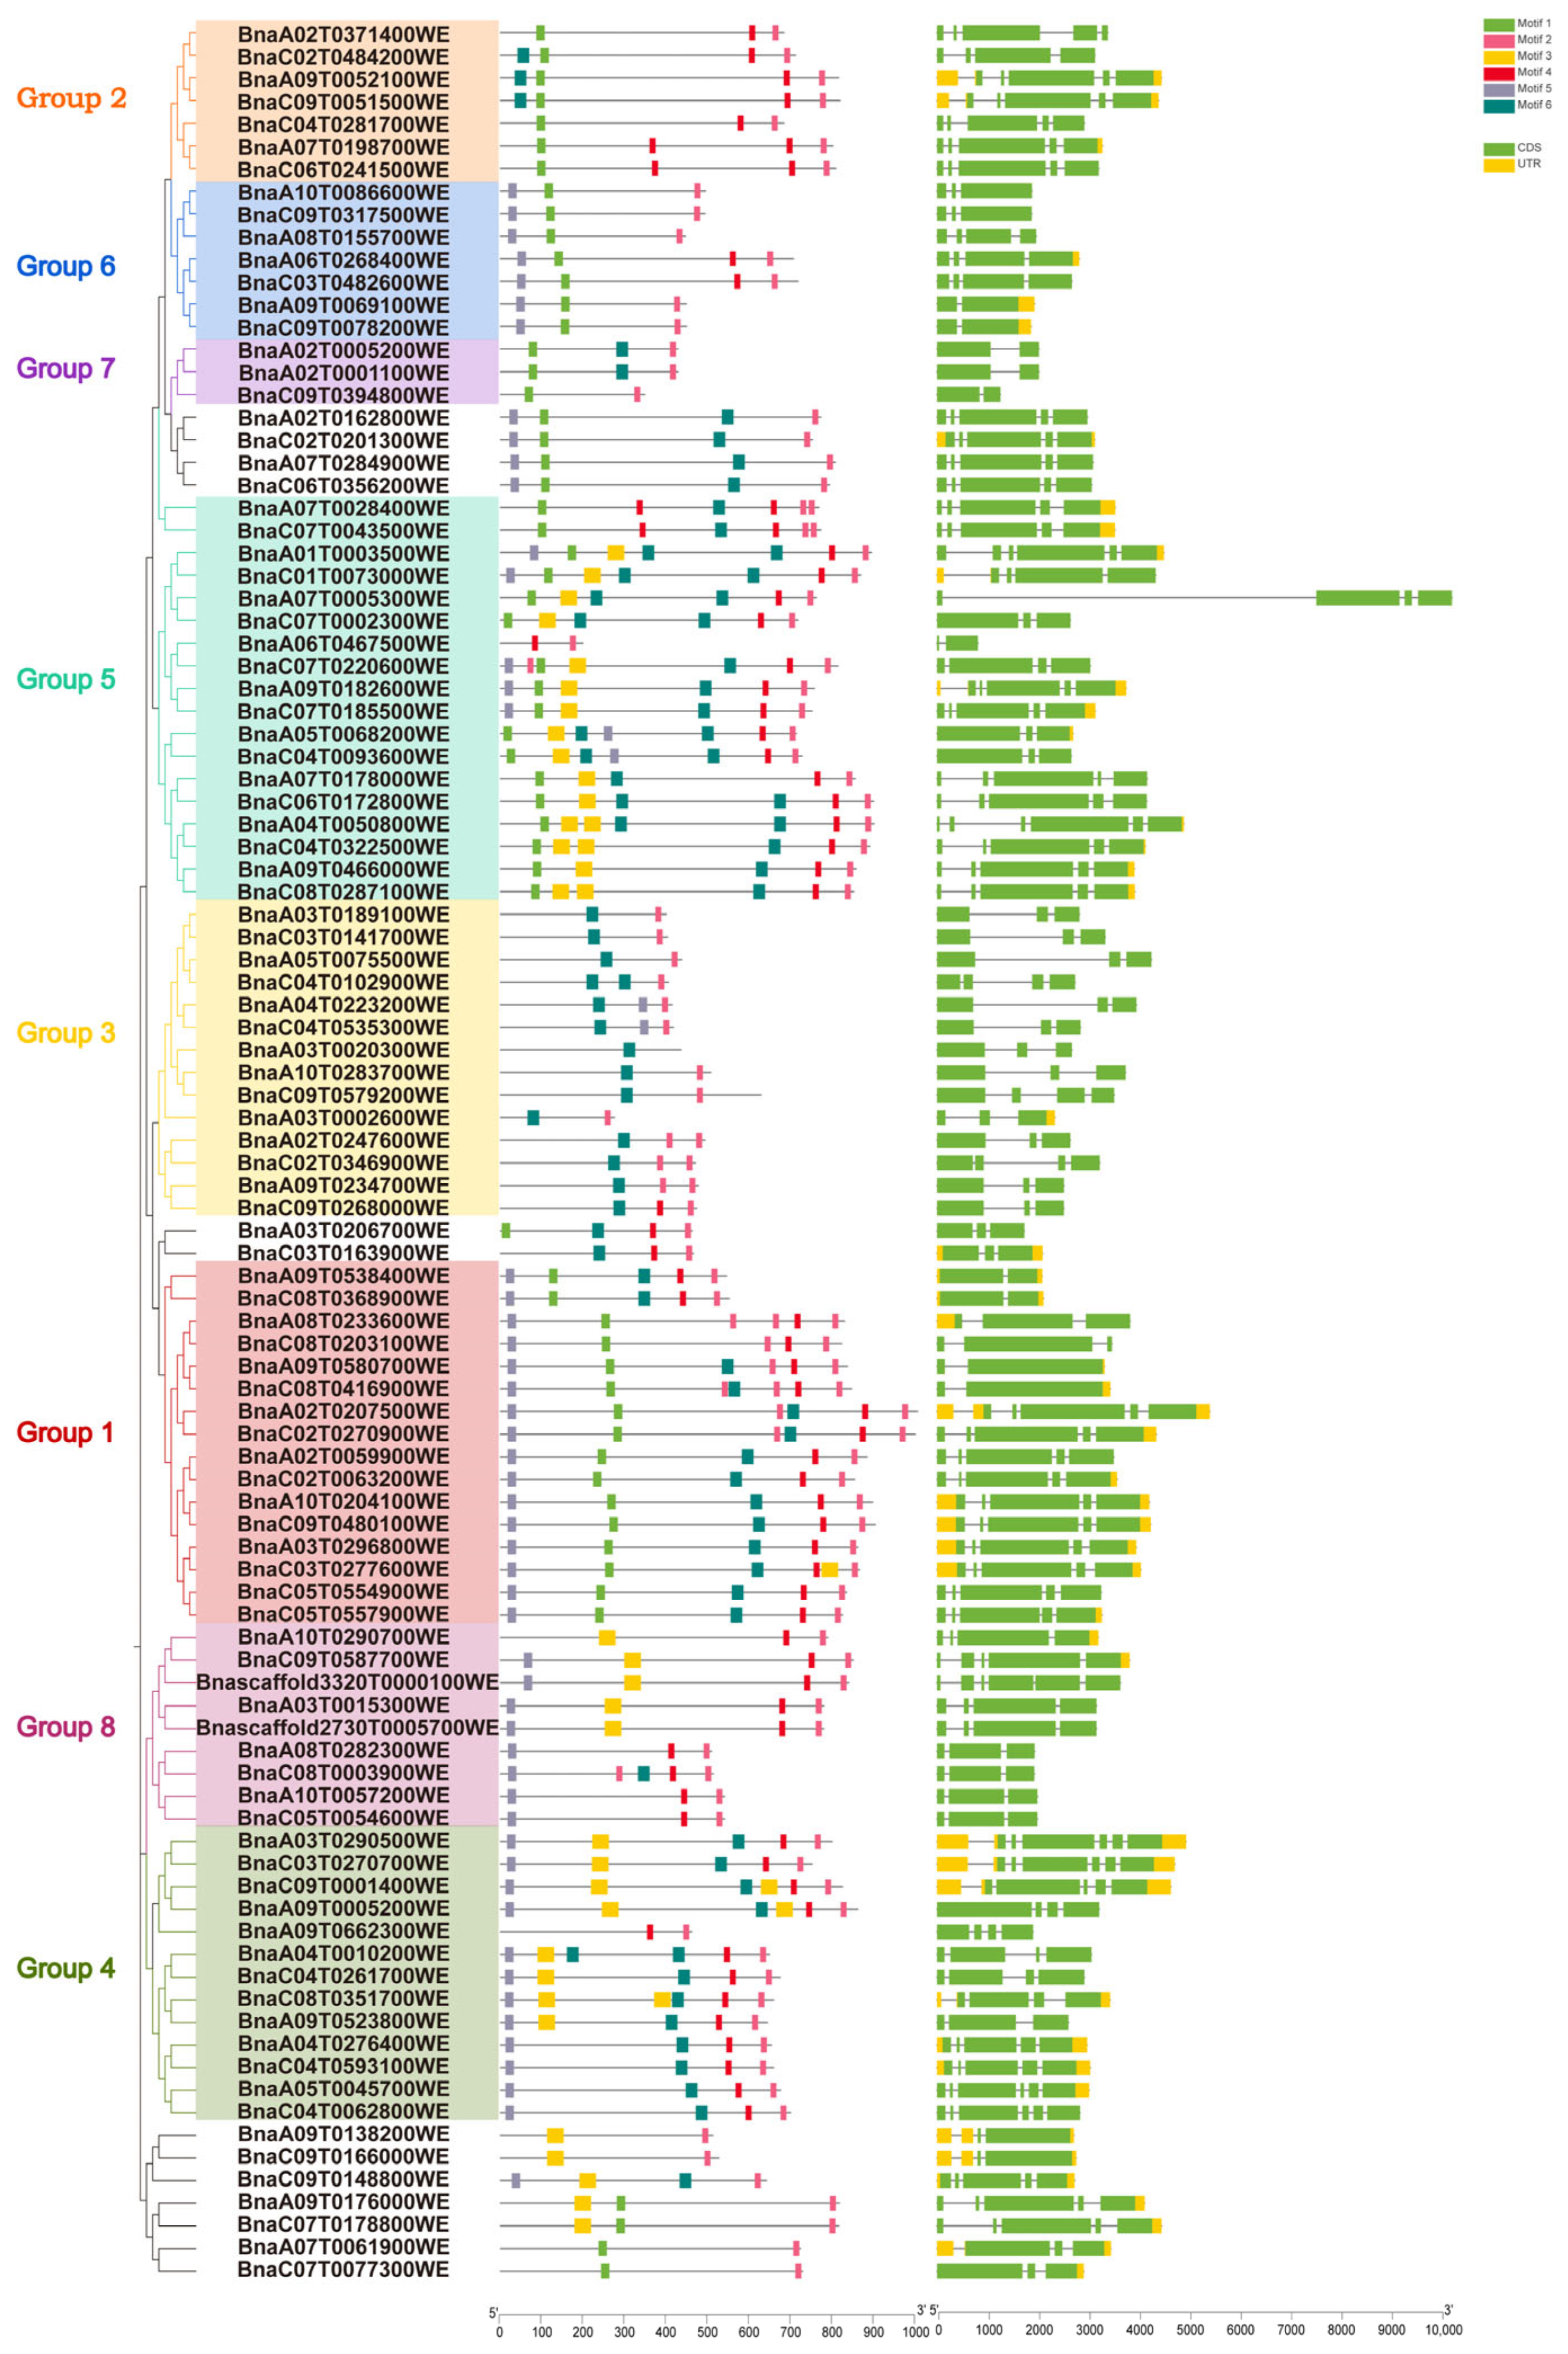
In Arabidopsis thaliana, 34 TRM proteins possess six highly conserved motifs, which are strictly conserved in the order of M5-M1-M3-M6-M4-M2 along the protein sequence. All 34 AtTRMs contain motif M2 at their C-termini, potentially serving as a characteristic feature of the entire TRM superfamily. Based on this feature, 100 BnaTRM genes conforming to this characteristic were selected from the 128 homologous genes, and their motifs were subjected to visual analysis (Figure 2). Some BnaTRM only possess two conserved motifs, while most of them own 4–5 (Figure 3). The M1-M6 motif in Brassica napus is slightly longer than the M1-M6 motif in Arabidopsis thaliana, but the sequence is relatively conserved. The conservation of the M2 motif ensures the stability of core functions such as microtubule nucleation [11], whereas the dynamic loss of variable motifs (e.g., M4) constitutes an evolutionary trade-off mechanism. This evolutionary strategy not only preserves the functional integrity of TRM proteins but also endows plants with environmental adaptability innovations through module replacement. This evolutionary paradigm exhibits significant convergence with the module recombination mechanisms observed in the TRM gene family of Cucumis sativus L. [30].
Figure 2.
Sequences of top six enriched motifs identified by MEME v5.5.7. The MEME v5.5.7 tool was used with the following parameters: protein; nostatus; mod anr; nmotifs 6; minsites 17; minw 10; maxw 100.
Figure 3.
Sequence logo representation of top eight enriched six motifs defined from the 100 BnaTRM proteins by MEME v5.5.7.
To further investigate the gene structure, a comparison was conducted between the CDS and genomic sequences of the BnaTRM genes. The comparison results showed that BnaA02T0005200WE, BnaA02T0001100WE, BnaC09T0394800WE, BnaA06T0467500WE, BnaA09T0538400WE, BnaC08T0368900WE, BnaA09T0580700WE, and BnaC08T0416900WE comprised only two exons. The remaining genes contained at least three exons (Figure 3). The difference in the number of exons reflects the hierarchical complexity of gene expression regulation: double-exon genes (such as BnaA02T0005200WE) may achieve tissue-specific expression through cis-elements in the promoter region (such as ABRE/ARE), while multi-exon genes are more likely to produce functional isomers through alternative splicing to meet the needs of different development stages.
2.3. Chromosome Distributions of the BnaTRM Genes
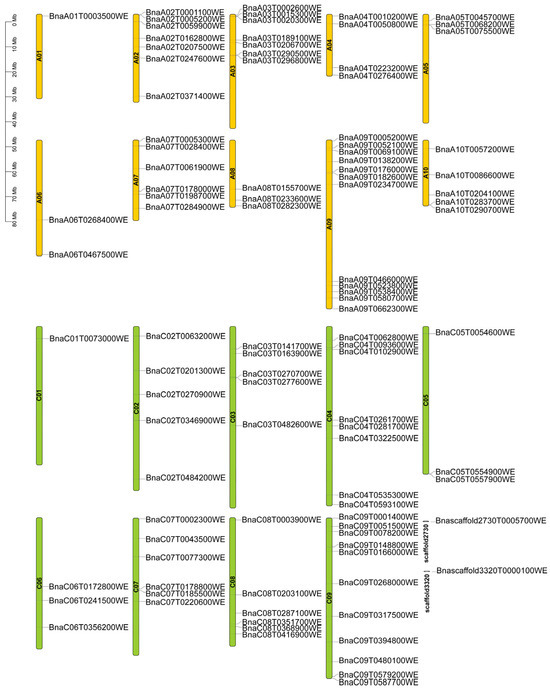
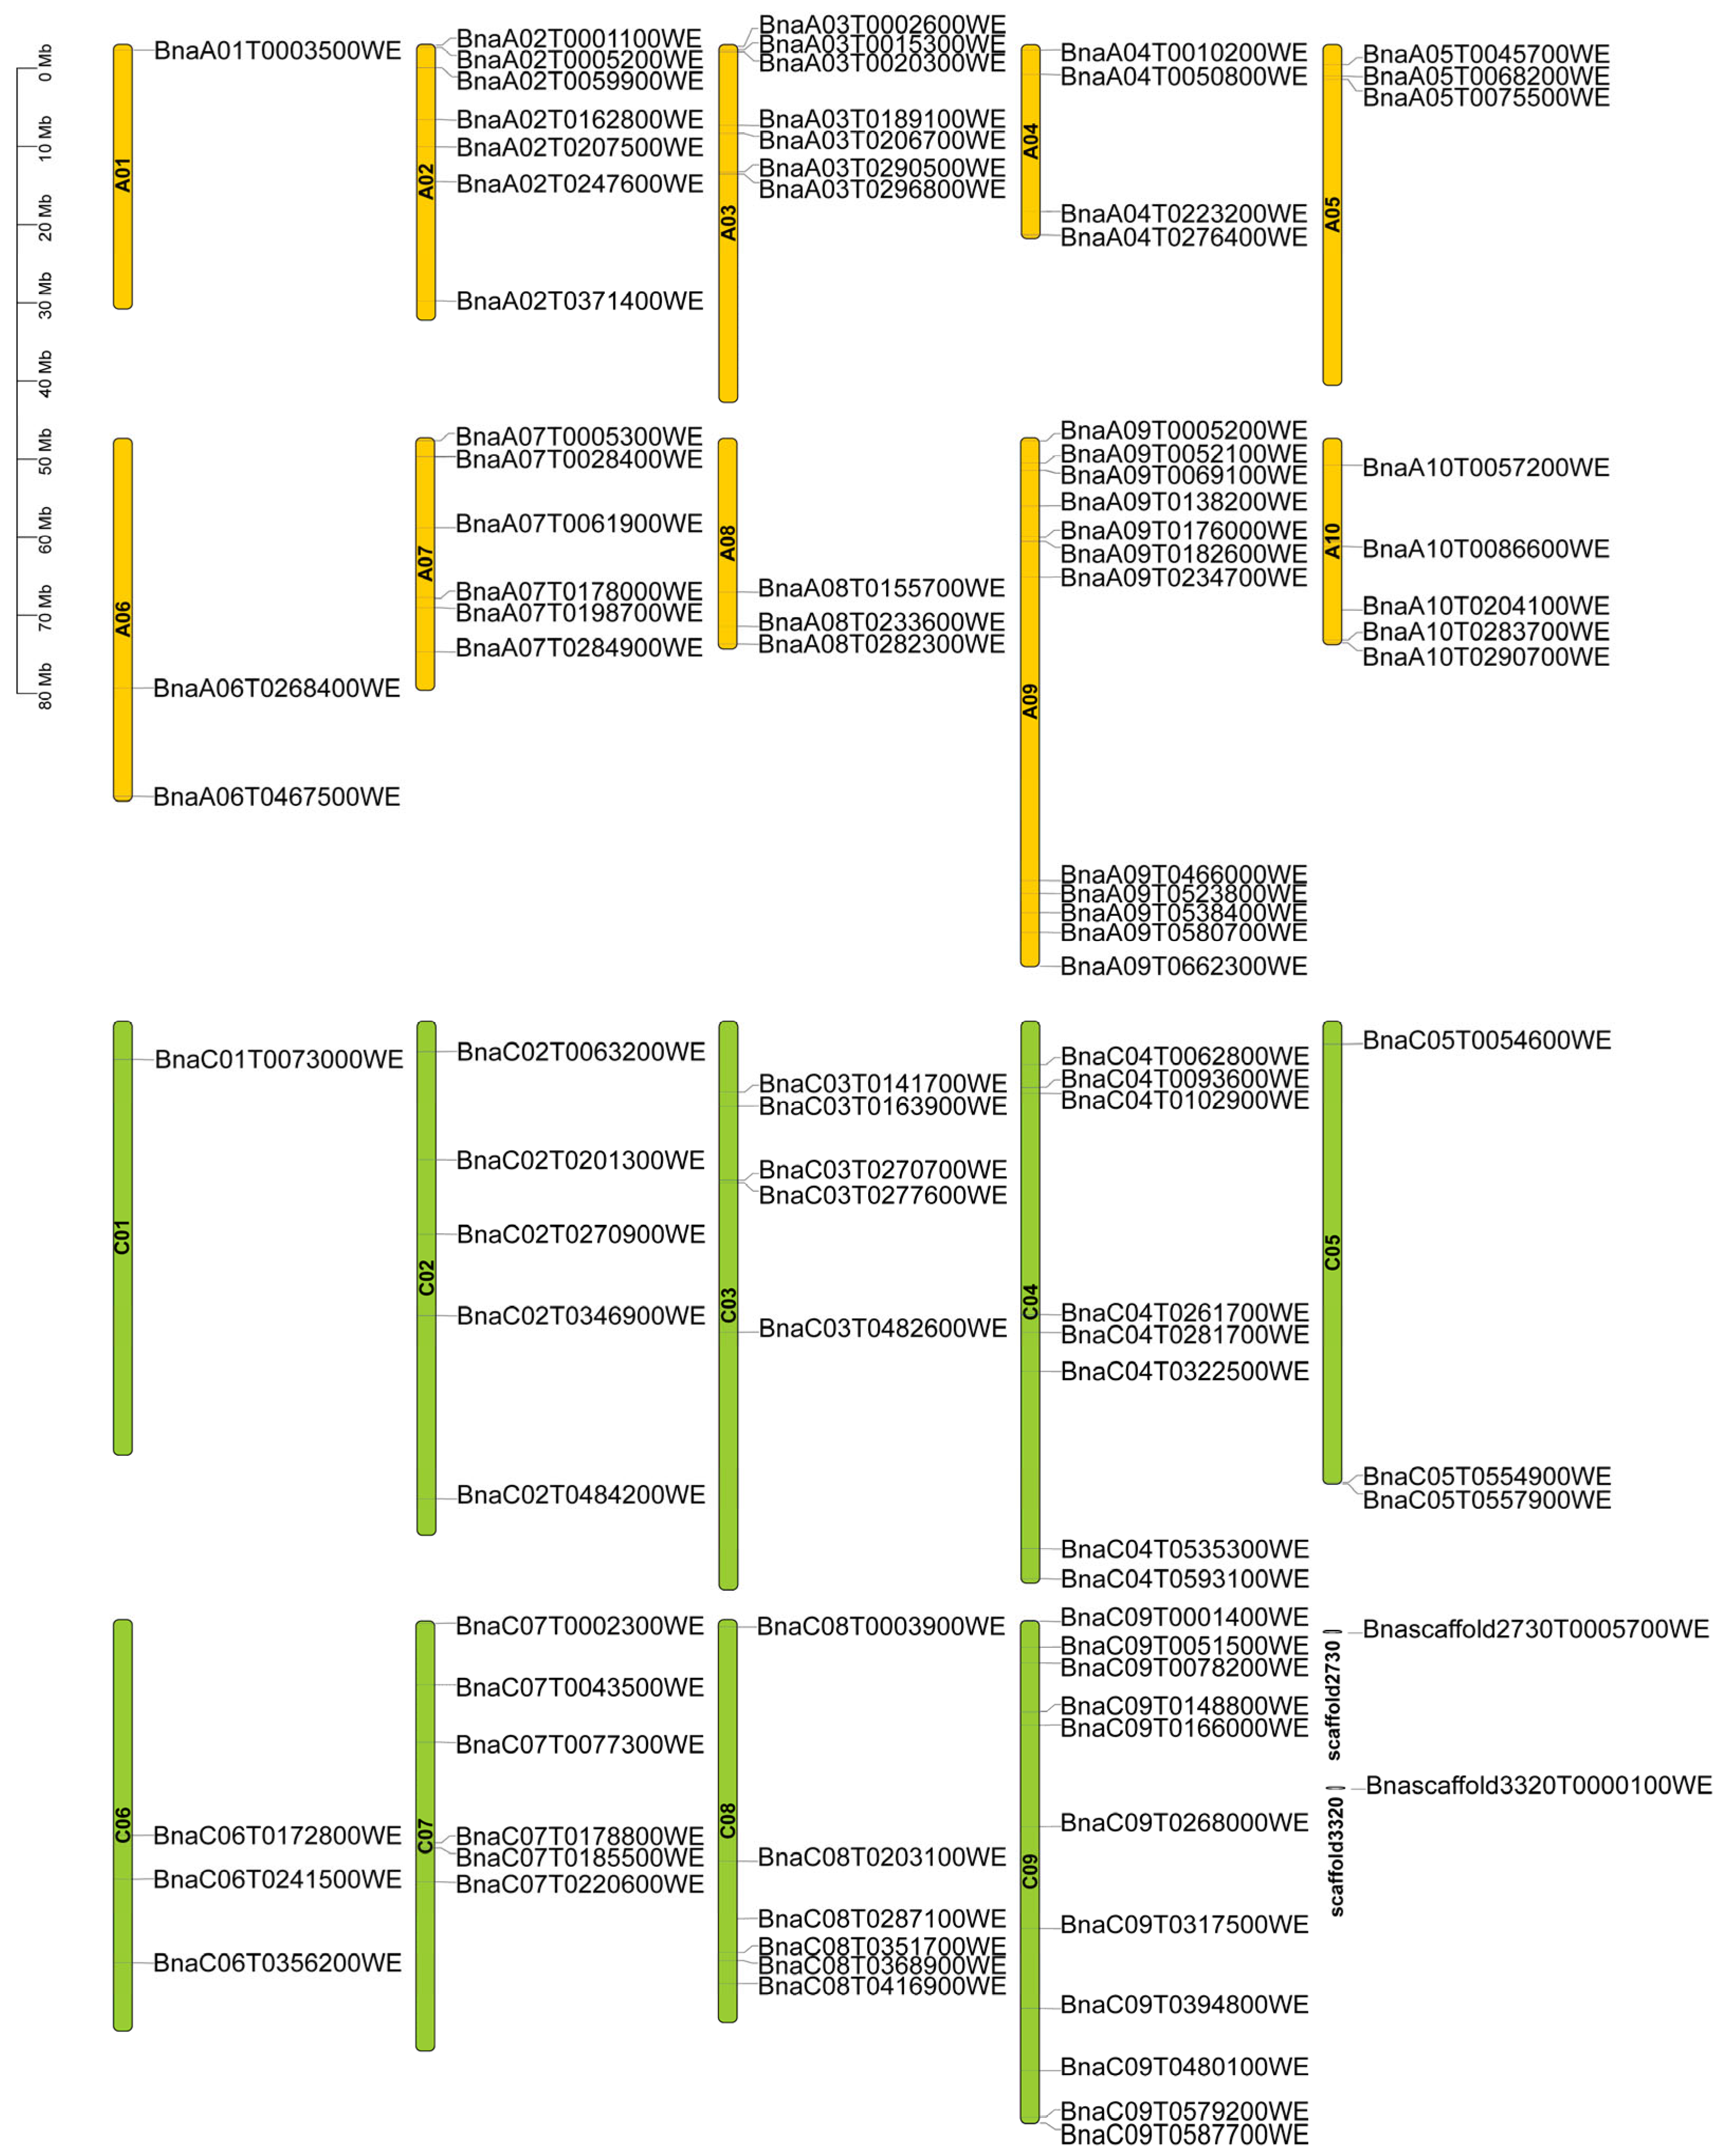
To gain a more intuitive understanding of the distribution of TRM genes across the chromosomes of Brassica napus, we utilized TBtools v2.310 to create a chromosomal distribution map (Figure 4). Among the 19 chromosomes of Westar, 98 BnaTRM genes are distributed. Specifically, 50 BnaTRMs are located in the A subgenome, and 48 BnaTRMs are located in the C subgenome. Two genes, namely Bnascaffold2730T0005700WE and Bnascaffold3320T0000100WE, could not be mapped to specific chromosomes in the Westar genome and are instead situated on scaffolds2730, respectively. Chromosomes A09 and C09 contain the highest number of BnaTRM genes, each containing 11, while chromosomes A01 and C01 each contain only one BnaTRM gene. It is suggested that A09 and C09 may be the core regulatory regions mediated by TRM. This distribution pattern reveals the adaptive evolutionary strategies of the TRM gene family following polyploidization in Brassica napus.
Figure 4.
Chromosome locations of BnaTRM family genes. The length of the bars indicates the sizes of Brassica napus chromosomes. The physical locations of the genes are labeled on the left of the chromosomes, and the genes are labeled on the right of the chromosomes.
2.4. Synteny Analysis of BnaTRM Genes
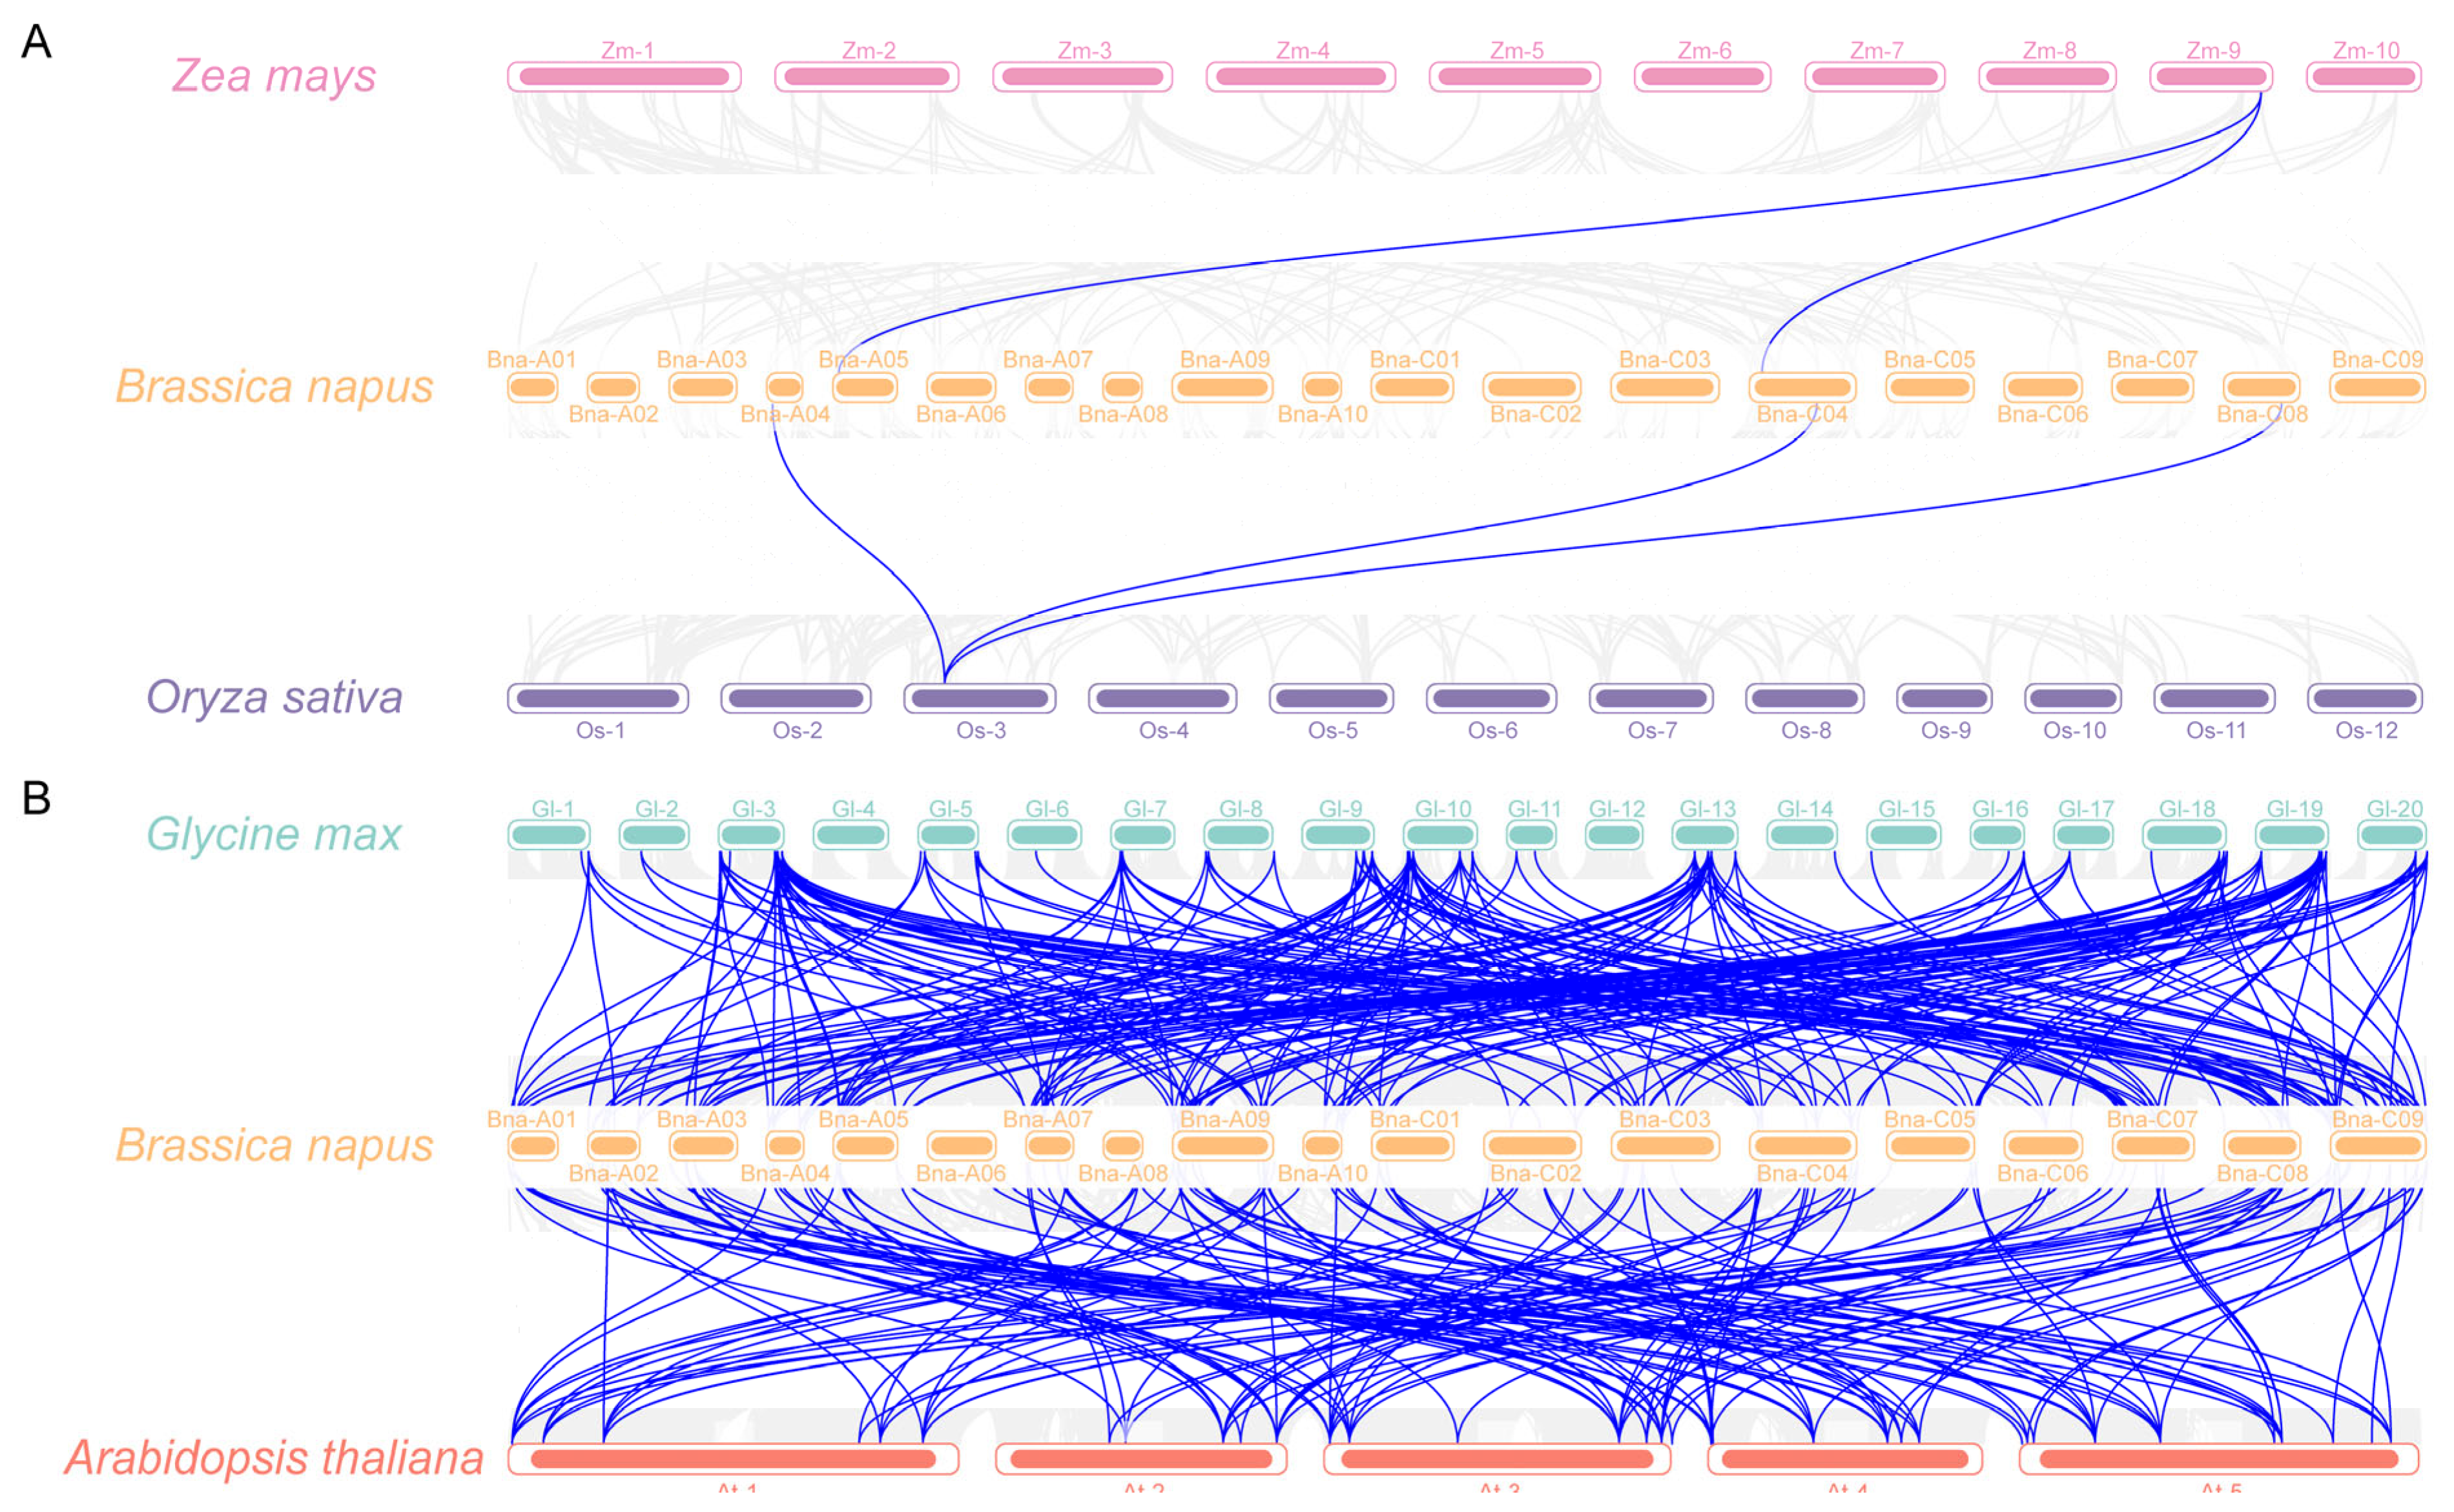
To delve deeper into the phylogenetic relationships of the Brassica napus TRM family, we constructed a gene collinearity map for Brassica napus and compared it with four representative species: two monocotyledonous plants (Oryza sativa L. and Zea mays, Figure 5A) and two dicotyledonous plants (Arabidopsis thaliana and Glycine max, Figure 5B). The results indicated that among Brassica napus and these four species, only 2 pairs of TRM collinear gene pairs were identified between Brassica napus and Zea mays, followed by 3 pairs between Brassica napus and Oryza sativa, 152 pairs between Brassica napus and Arabidopsis thaliana, and 224 pairs between Brassica napus and Glycine max. Notably, the number of orthologous genes shared between dicotyledonous plants was significantly higher than that shared between dicotyledonous and monocotyledonous plants. This observation aligns with the expected pattern of biological evolution. The collinear gene pairs identified between Brassica napus and Oryza sativa, Zea mays, and Arabidopsis thaliana were also present in Brassica napus and Glycine max, suggesting a possible shared evolutionary history between Brassica napus and Glycine max.
Figure 5.
Analysis of syntenic relationships among Oryza sativa, Zea mays L. (A), Arabidopsis thaliana, and Glycine max (B). The gray line represents the syntenic block in plant genomes, and the blue line represents the collinear TRM gene pair.
2.5. The Biophysical Properties of BnaTRMs
The molecular weights of the BnaTRM proteins ranged from 22.545 kDa (BnaA06T0467500WE) to 111.547 kDa (BnaA02T0207500WE) in Brassica napus. The pI ranged from 4.37 (BnaA03T0002600WE) to 9.86 (BnaC09T0078200WE). On the basis of their pI, most BnaTRM proteins were acidic. Fifty-eight proteins are acidic, and thirty-seven proteins are alkaline. However, all of the BnaTRM proteins were unstable, with an instability index greater than 40. The Grand Average of the Hydropathicity index for all TRM proteins (<0) reflected the hydrophilic nature of these proteins. The aliphatic index for all BnaTRM proteins ranged from 59.27 (BnaA05T0075500WE) to 80.14 (BnaA09T0538400WE). The subcellular localization analysis revealed that the BnaTRM proteins were mostly localized in the nucleus (87 BnaTRMs), followed by the chloroplast (6 BnaTRMs) and cytoplasm (2 BnaTRMs) (Table 1). This analysis reveals the evolutionary strategy of the BnaTRM family to realize functional specialization through physical and chemical property differentiation.
2.6. Cis-Regulatory Elements Identification in Promoters
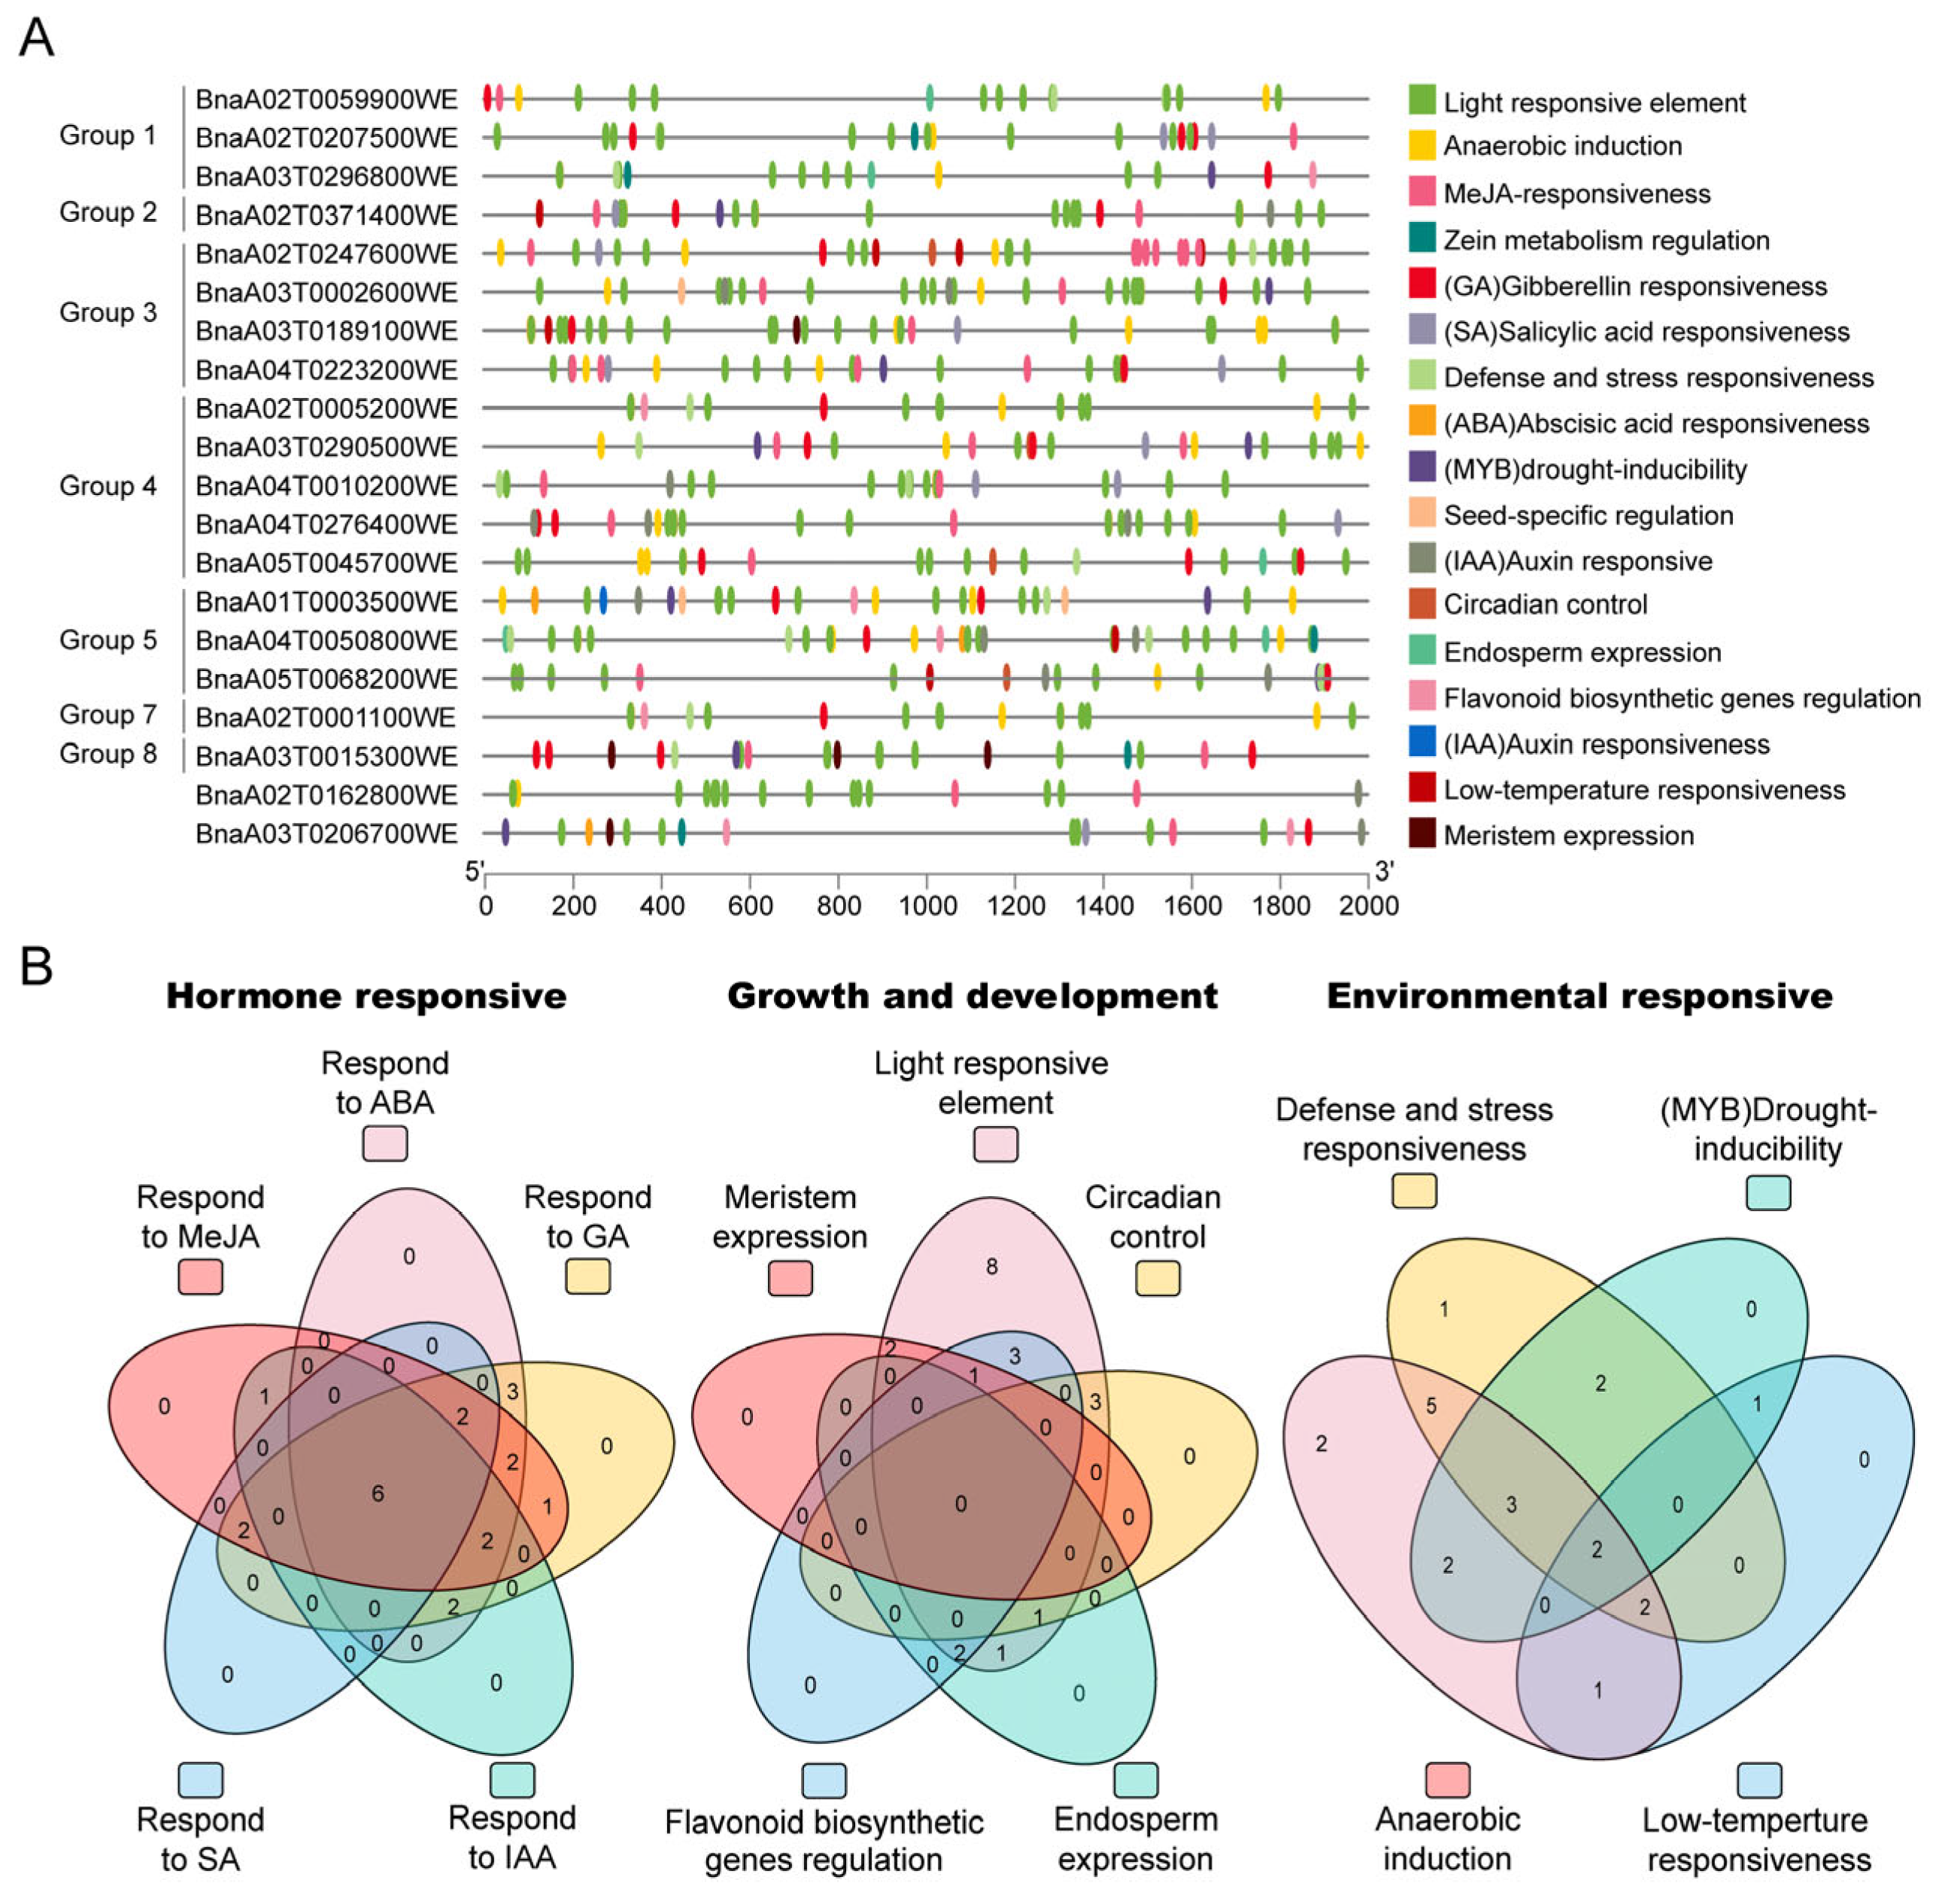
Cis-acting elements in the promoter region are very important for regulating the expression of corresponding genes because they bind to various transcription factors. Thus, we identified the cis-regulatory elements in 2000 bp sequences upstream of the start codon of genes in BnaTRM using the PlantCARE database. As predicted, common promoter elements (TATA-box and CAAT-box) were identified among all BnaTRM gene promoters. Among 526 cis-acting elements, 233 cis-elements related to light, 172 cis-elements related to hormones, and 121 cis-elements with other functions were detected. Among the cis-acting elements related to hormones, 68, 42, 31, 18, and 13 are involved in MeJA, ABA, GA, IAA, and SA responsiveness. Therefore, BnaTRMs have the potential to play a role in a variety of processes. Forty-six light-responsive cis-elements involved in growth and development were detected in the TRM gene promoter regions (Table 2). Among the cis-acting elements involved in hormone responses, ABRE, GAREs (GARE-motif and P-box), and the MeJA-responsive elements (CGTCA-motif and TGACG-motif) were, respectively, identified in the promoter regions of 17, 19, and 16 TRM genes in Brassica napus. The zein metabolism regulation element (O2 site) was detected in six BnaTRM genes, whereas the CAT-box influencing meristem expression was identified in the promoter regions of three BnaTRM genes. Of the stress-related response elements, ARE, which is an anaerobic induction element, was detected in the promoters of 17 TRM genes. Some stress-related (low temperatures, wounding, and drought) cis-acting elements were also identified in the promoter regions of TRM genes. We use TBtools v2.310 to visualize these data, as shown in Figure 6. This analysis reveals the evolutionary strategy of the BnaTRM gene family to realize multi-dimensional environmental response through a modular combination of cis-acting elements, and the related functions of BnaTRM with core regulatory elements (such as ABRE and GAREs) can be verified first in the future.
Table 2.
Number of hormone and stress-responsive cis-elements in the promoters of the BnaTRMs.
Figure 6.
Cis-elements analysis in the promoter sequences of BnaTRM genes: (A) Visualization of the cis components of the TRM family with TBtools v2.310. (B) The Venn diagram is used to show the cis components under different paths.
2.7. Expression Profiling Analysis of 29 BnaTRM Genes Across 12 Distinct Tissues
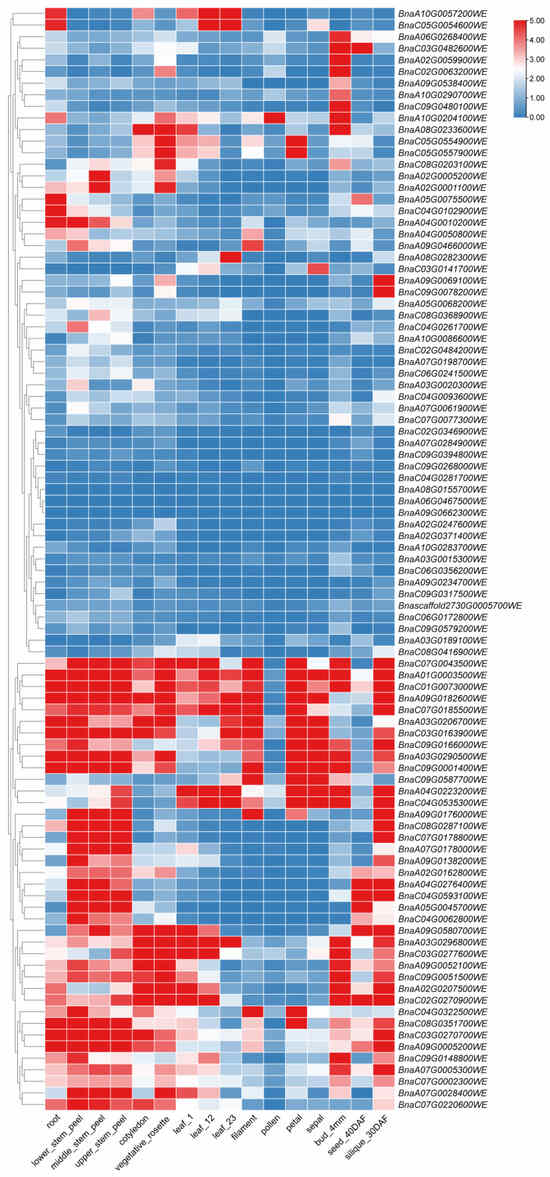
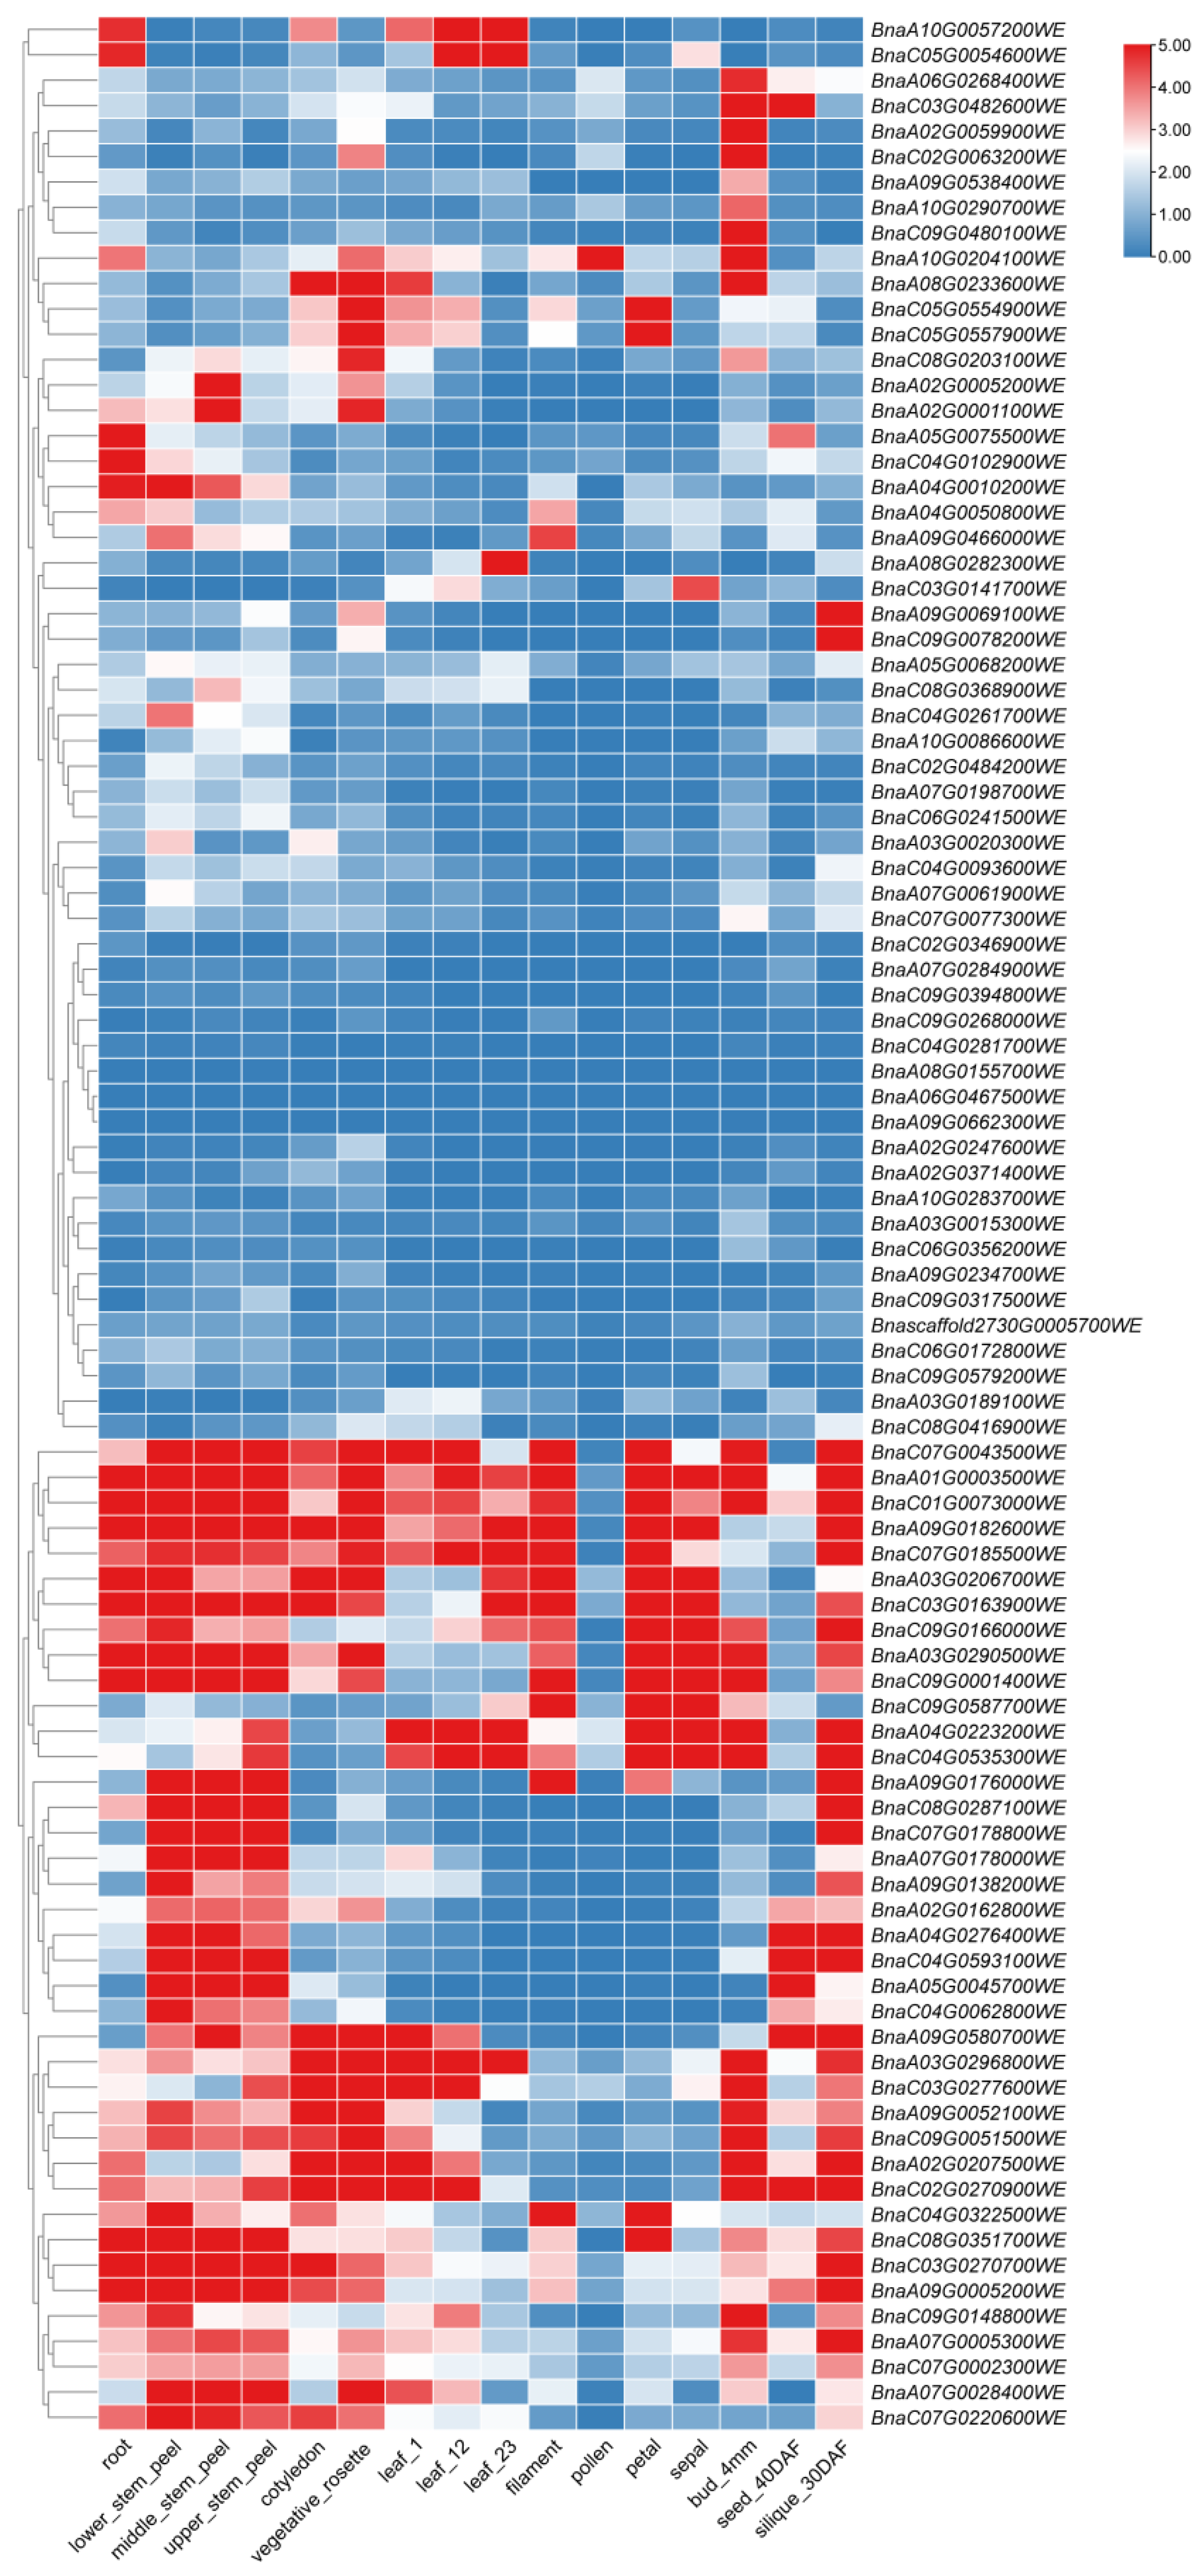
Gene expression levels are typically closely associated with their biological functions. To investigate the functional roles of the BnaTRM genes family in Brassica napus, this study utilized the publicly available transcriptome database of the Brassica napus cultivar Zhongshuang 11 (ZS11) (https://yanglab.hzau.edu.cn/BnIR/expression_zs11 (accessed on 5 March 2024)). The BnaTRM genes family members from the Westar cultivar were homologously mapped to the ZS11 cultivar, and the expression patterns of BnaTRM genes across 12 different tissues (root, stem, cotyledon, vegetative rosette, cauline leaf, filament, pollen, petal, sepal, bud, seed, and silique) were analyzed using TBtools v2.310 software (Figure 7). The key findings revealed that most BnaTRM genes were upregulated in roots and stems. Five genes belonging to the Group 5 subfamily (BnaC07T0043500WE, BnaA01T0003500WE, BnaC01T0073000WE, BnaA09T0182600WE, BnaC07T0185500WE) exhibited low expression levels in pollen and seeds but showed high expression in the other 10 tissues. Eight genes from the Group 3 subfamily (BnaC02T0346900WE, BnaC09T0268000WE, BnaA02T0247600WE, BnaA03T0189100WE, BnaA09T0234700WE, BnaA10T0283700WE, BnaC09T0579200WE, BnaA03T0020300WE) displayed low expression across all tissues. These genes may regulate critical pathways in seed development (e.g., lipid biosynthesis or dormancy regulation), warranting further investigation into their association with agronomic traits such as seed size and oil content. The tissue-specific expression patterns of BnaTRM genes suggest that they are implicated in multiple regulatory processes during plant growth and development.
Figure 7.
Expression patterns of TRM family genes in twelve tissues. The heat map was generated using log2 expression levels (TPM + 1). The bar indicates the log2 expression levels (TPM + 1). The legend label is shown on the right side of the figure. Genes with high expression levels are shown in red, and genes with ground expression levels are shown in blue. No dates are available for the BnaA03T0002600WE, BnaA09T0523800WE, BnaC02T0201300WE, BnaC08T0003900WE, and Bnascaffold3320T0000100WE genes. The RNA-seq data comes from the website BNIR (BnIR, Brassica napus multi-omics database (information resource).
2.8. Differential Expression Analysis of BnaTRM Gene Family in Cold-Tolerant Varieties and Cold-Sensitive Varieties
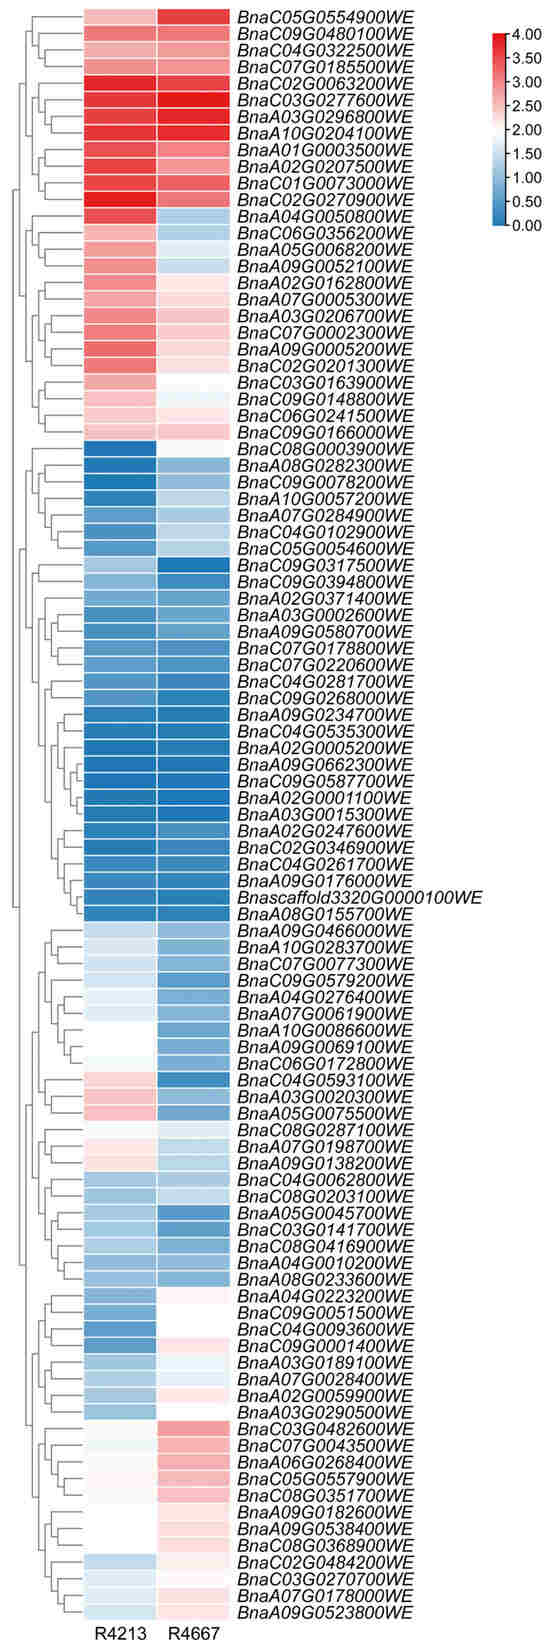
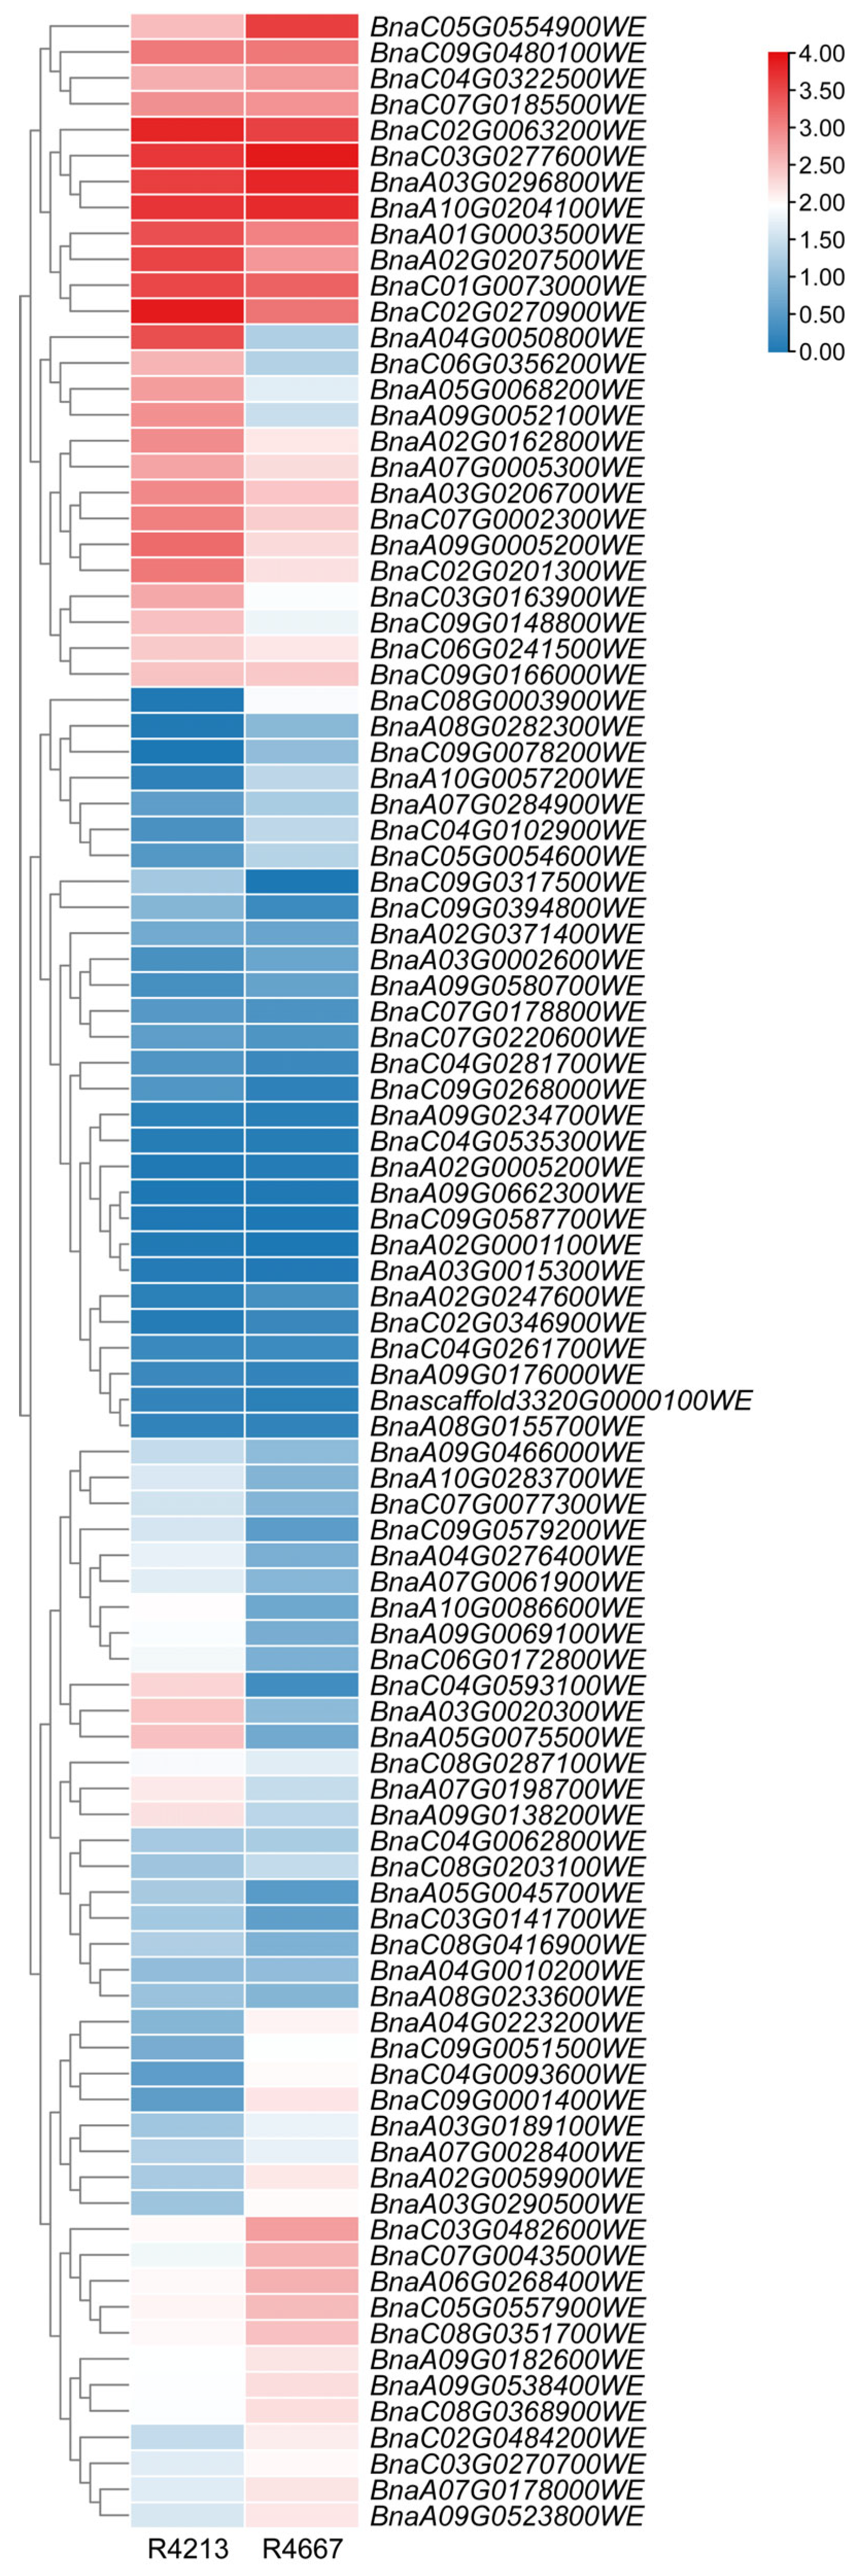
Based on the method of cold-tolerant field-grown cultivar identification [37,38], we used the inflorescence materials of one cold-tolerant R4213 and another cold-sensitive R4667 cultivar for RNA-seq profiling. The heat map demonstrated significant divergence in BnaTRM expression profiles between the two groups (Figure 8). In cold-tolerant varieties, 31 genes such as BnaA02T0207500WE, BnaA05T0068200WE, and BnaA02T0162800WE showed remarkable upregulated patterns with extremely significant differential expression compared to the cold-sensitive varieties. These genes play a positive role in regulating cold stress. On the contrary, the expression levels of 23 genes, such as BnaA02T0059900WE, BnaC05T0557900WE, and BnaA10T0057200WE, in cold-tolerant varieties were significantly lower than those in cold-sensitive varieties, which may play a negative regulatory role in cold resistance. More than half of BnaTRM genes may respond to cold stress, indicating that BnaTRM genes may contribute to the low-temperature adaptation of Brassica napus.
Figure 8.
Expression patterns of TRM family genes in main inflorescence in cold-tolerant and cold-sensitive cultivars grown in the field. The heat map was generated using log2 expression levels (TPM + 1). The bars indicate the log2 expression levels (TPM + 1). The color scales represent relative expression levels from high (red color) to low (blue color). No dates are available for the Bnascaffold2730G0005700WE, BnaA06G0467500WE, and BnaC04G0261700WE genes.
2.9. Transcriptional Response of TRM Genes to Abiotic Stress in Brassica napus
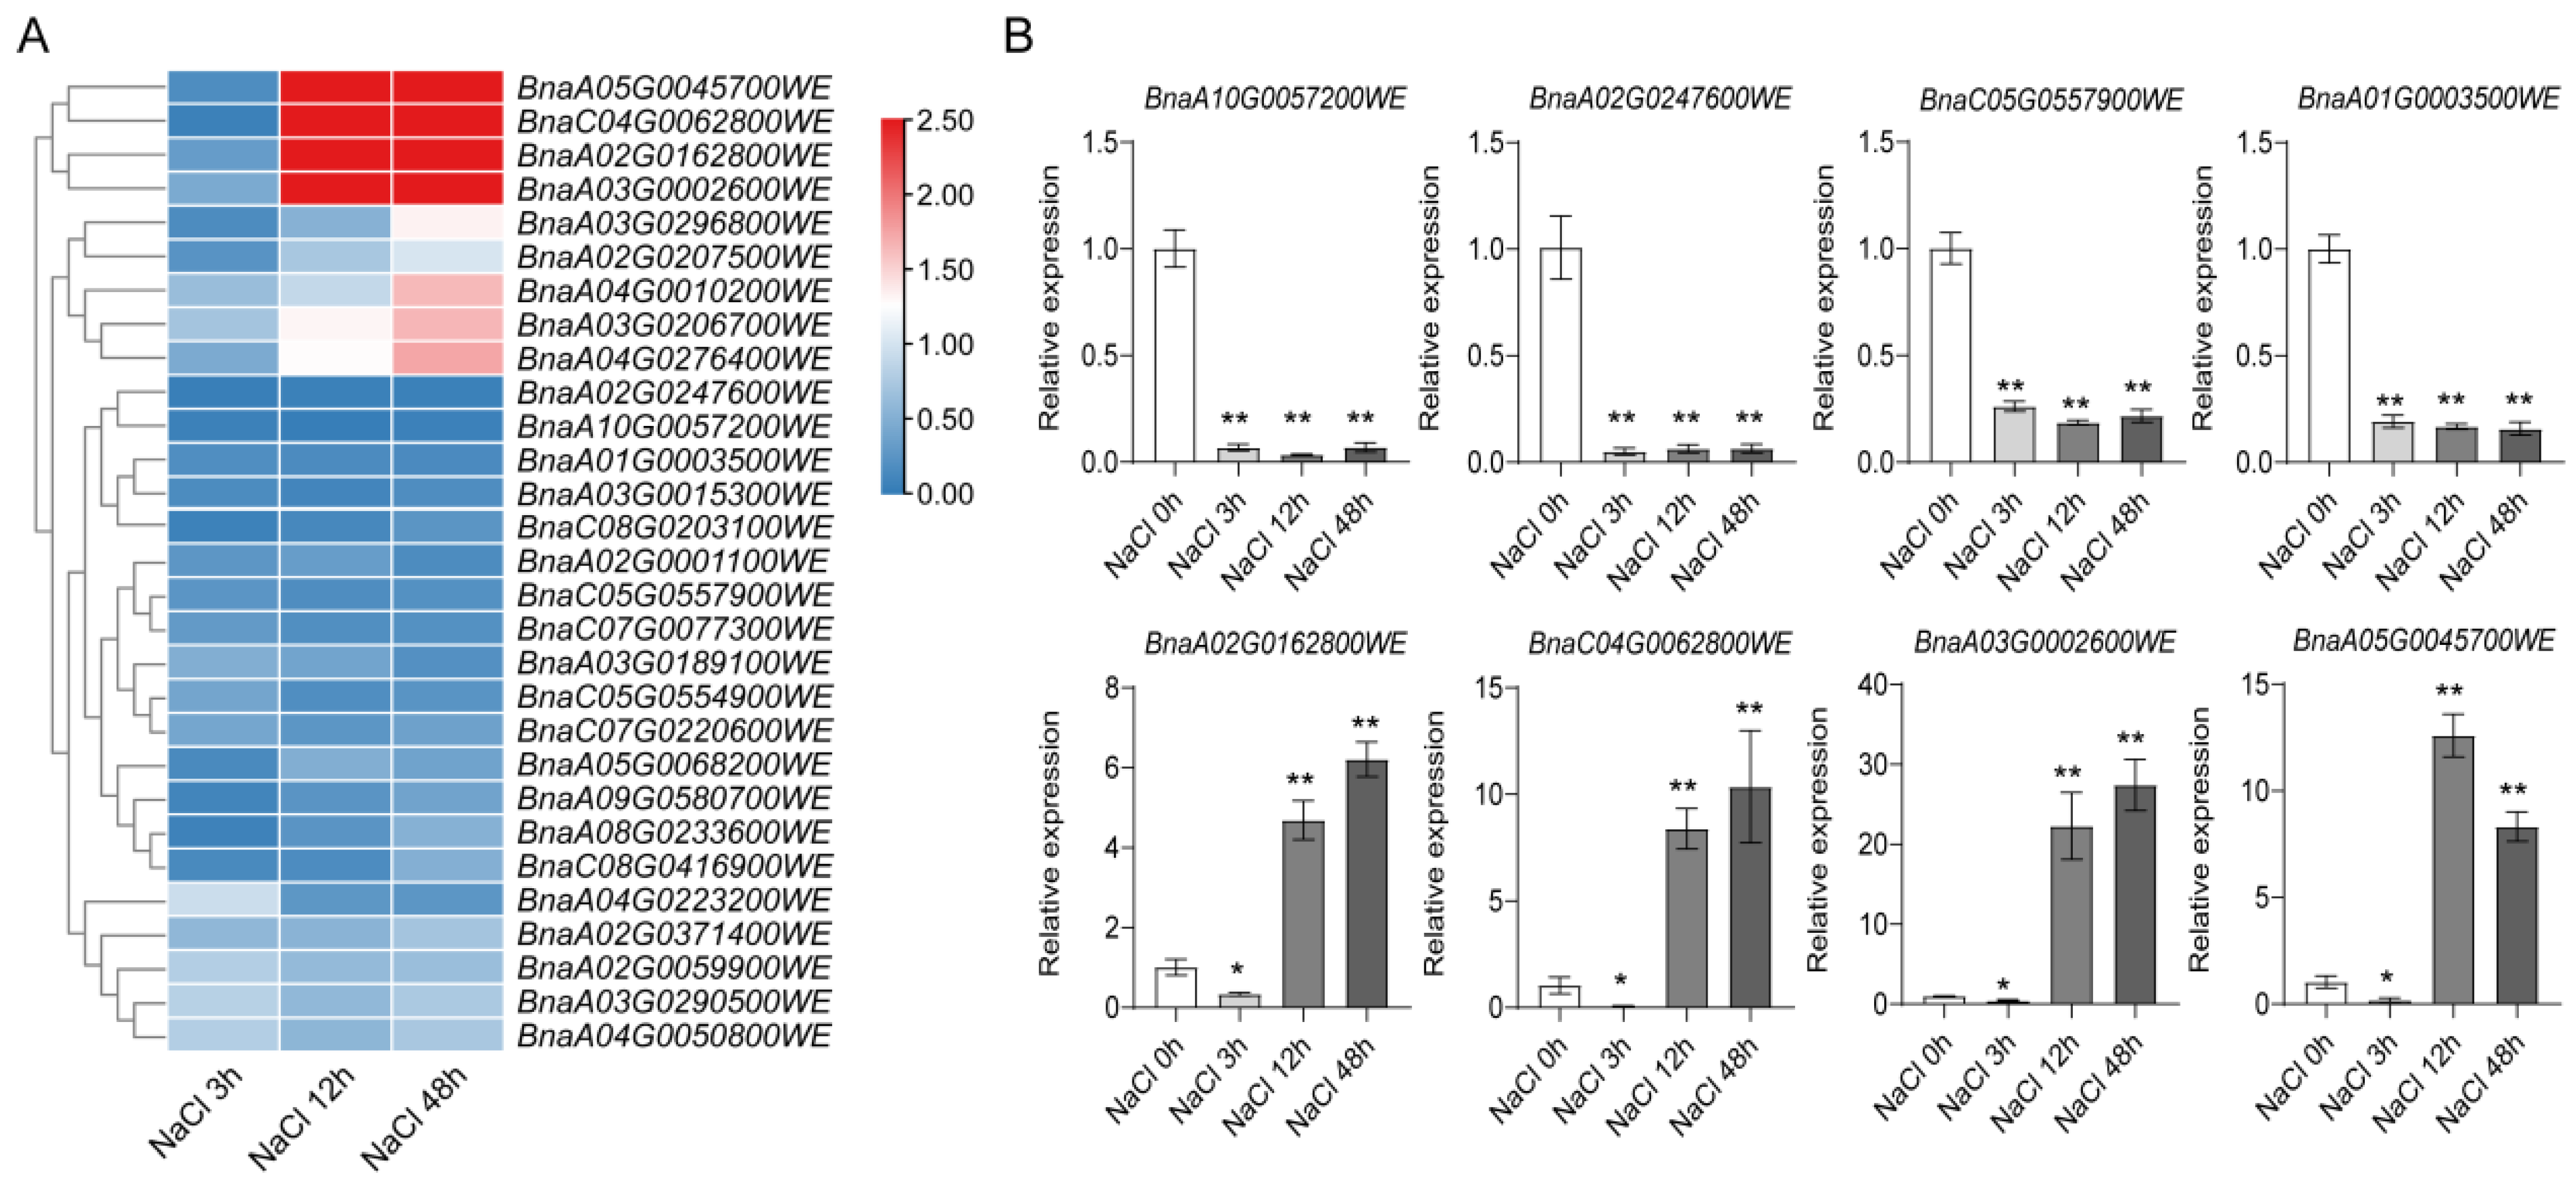
Abiotic stresses significantly impact plant growth and development. To investigate the effects of drought and salt stress on TRM gene expression in Brassica napus, we analyzed the transcriptional responses of 29 TRM genes under stress conditions using qRT-PCR. These 29 TRM genes include 20 genes containing cis-element, and 9 genes are homologous to the genes that are known to be involved in stress [30] (Figure 9A and Figure 10A). Under salt treatment, all TRM genes showed downregulated expression at 3 h post-treatment. Notably, four genes (BnaA10T0057200WE, BnaA02T0247600WE, BnaC05T0557900WE, and BnaA01T0003500WE) exhibited significantly reduced expression compared to controls. Interestingly, eight genes (BnaA05T0045700WE, BnaC04T0062800WE, BnaA02T0162800WE, BnaA03T0002600WE, BnaA03T0206700WE, BnaA04T0276400WE, BnaA03T0296800WE, and BnaA04T0010200WE) displayed unique dynamic regulation patterns, with significant upregulation observed at 12 h post-salt treatment (Figure 9B).
Figure 9.
Expression level of TRM family genes after salt (NaCl) treatment at different times. (A) The heat map was generated using log2 expression levels (TPM + 1). The bars indicate the log2 expression levels (TPM + 1). The color scales represent relative expression levels from high (red color) to low (blue color). (B) The relative expression analysis of the BnaTRMs under salt (NaCl) treatment. The x-axis presents the treatments. The y-axis presents the expression levels relative to the expression at the 0 h time point. The data are representative of three independent experiments (n = 3, mean ± SD, * p < 0.05, ** p < 0.01, Student’s t-test).
Figure 10.
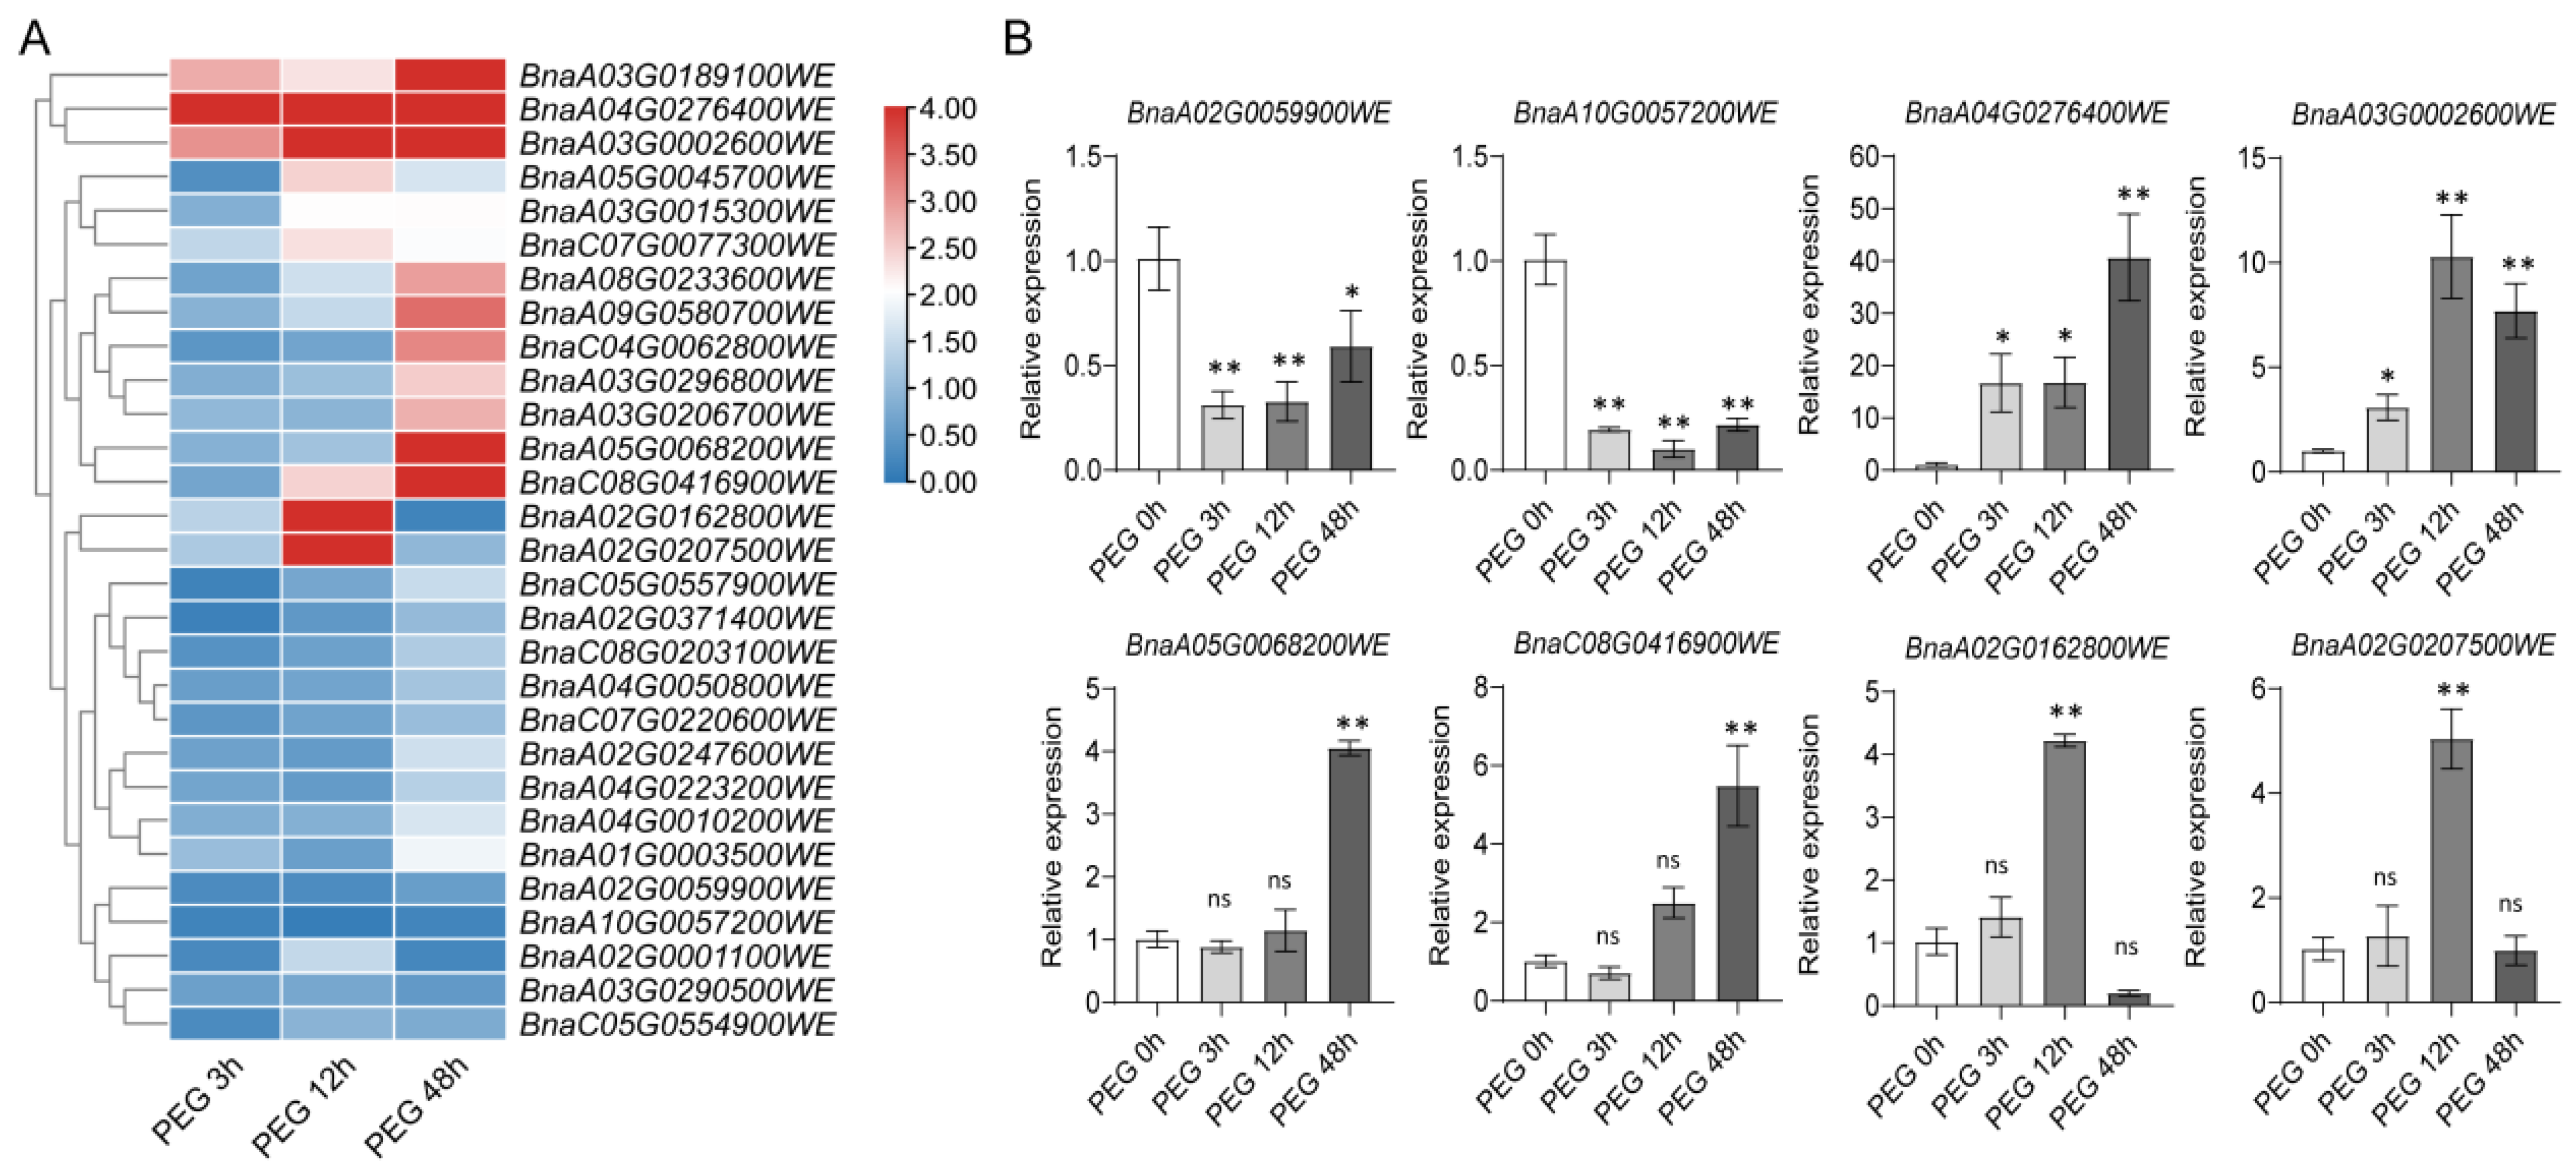
Expression level of TRM family genes after drought (PEG) treatment at different times: (A) The heat map was generated using log2 expression levels (TPM + 1). The bars indicate the log2 expression levels (TPM + 1). The color scales represent relative expression levels from high (red color) to low (blue color). (B) The relative expression analysis of the BnaTRMs under drought (PEG) treatment. The x-axis presents the treatments. The y-axis presents the expression levels relative to the expression at the 0 h time point. The data are representative of three independent experiments (n = 3, mean ± SD, * p < 0.05, ** p < 0.01, Student’s t-test) and ns represents no significant difference.
Meanwhile, after 3 h of drought treatment, we observed distinct transcriptional responses among TRM genes: BnaA02T0059900WE and BnaA10T0057200WE exhibited significant downregulation, while BnaA04T0276400WE and BnaA03T0002600WE showed remarkable upregulation with extremely significant differential expression (fold change ≥ 10) compared to controls. Notably, BnaA05T0068200WE and BnaC08T0416900WE displayed delayed response patterns, with significant upregulation only observed after 48 h of drought treatment. In contrast, BnaA02T0162800WE and BnaA02T0207500WE demonstrated biphasic regulation, showing upregulated expression at 12 h post-treatment, but they underwent significant downregulation by 48 h (Figure 10B). It is worth noting that BnaA03T0002600WE showed an upregulation under both drought and salt stress. It was demonstrated that this gene was positively involved in the stress response. However, BnaA10T0057200WE showed a continuous decrease under salt stress and drought stress, which may reflect its negative regulatory role in response to stress.
3. Discussion
TRMs are the TON1 Recruiting Motif (TRM) proteins, which share six short conserved motifs, including a TON1-interacting motif present in all TRMs. Members of the TRM protein family are generally positioned on microtubules and play a pivotal role in the establishment and preservation of microtubule architecture. Microtubules constitute a fundamental element of the plant cytoskeleton, essential for sustaining cellular shape, accommodating growth and development, and responding to environmental fluctuations. Furthermore, they are integral to processes such as cell division, material transportation, immune responses, and stress tolerance. Furthermore, it also contributes to regulating resistance to drought stress and plays an important role in plant growth and development.
In this study, we identified 100 TRM genes in Brassica napus, which is much higher than other species such as Oryza sativa, Glycine max, Triticum aestivum, and Cucumis sativus, perhaps because of complex multiple genome-wide duplication events in Brassica napus [39,40,41,42]. We used bioinformatics technology to conduct phylogenetic, collinearity, homology, gene structure, motif, chromosome location, expression, and cis-regulatory elements in promoter regions analyses for the TRM gene family members of Brassica napus. From this comprehensive gene family investigation, we drew three conclusions.
3.1. Genomic Architecture and Evolutionary Dynamics of TRM Genes in Brassica napus
The identification of 100 BnaTRM genes in Brassica napus reveals a complex genomic landscape shaped by evolutionary forces. The uneven distribution of BnaTRM genes across 19 chromosomes (with chromosomes A09 and C09 harboring 11 genes each) suggests functional clustering or tandem duplication events. This clustering may reflect the coordinated regulation of microtubule dynamics during specific developmental stages or stress responses. Comparative genomic analysis highlights the close evolutionary relationship between Brassica napus and Glycine max, with 224 collinear gene pairs identified. This synteny aligns with their shared Brassicaceae ancestry and supports the hypothesis of a common TRM gene family expansion in the Brassiceae tribe. In contrast, fewer orthologs were detected in monocots (Oryza sativa L. and Zea mays), indicating divergent evolutionary trajectories after the divergence of eudicots and monocots. Phylogenetic clustering into eight subfamilies (Groups 1–8) further underscores functional diversification, with Group 3 (18 genes) emerging as the largest clade.
3.2. Functional Prediction of BnaTRM Gene Family
Through comprehensive integrative analysis of the BnaTRM gene family (including cis-regulatory element identification, subcellular localization prediction, expression profiling, and quantitative validation), we systematically elucidated the multi-dimensional regulatory mechanisms of this gene family in plant stress responses. Promoter analysis revealed that BnaTRM genes are enriched with various stress-responsive elements (ARE, DRE, LTR) and hormone-responsive motifs (ABRE, MeJA-responsive CGTCA motif). Among them, 42 members harbor ABRE elements, suggesting their involvement in low-temperature, drought, and salt stress response pathways; 16 genes containing CGTCA motifs may mediate jasmonic acid signaling-mediated defense responses, while 17 ARE-containing genes potentially regulate anaerobic stress adaptation. Subcellular localization features indicate that nuclear-localized members (e.g., BnaA03T0002600WE) likely regulate downstream transcriptional networks under stress conditions through their ABRE/CGTCA-enriched promoters, whereas chloroplast-localized isoforms (e.g., BnaA09T0052100WE) imply TRM proteins may participate in photosynthesis or stress metabolic pathways by modulating cytoskeletal dynamics.
QRT-PCR validation further confirmed these predictions: BnaA03T0002600WE showed significant upregulation under drought and salt stress with vascular tissue-specific expression, suggesting its role in vascular system stress signaling transduction. Gene BnaA02T0162800WE exhibited consistently elevated expression under low-temperature, drought, and salt stresses, while BnaA10T0057200WE displayed downregulation across these conditions, indicating their divergent regulatory roles in plant abiotic stress adaptation. BnaA02T0162800WE may function as a positive regulator by modulating osmolyte accumulation, enhancing antioxidant defenses, or activating stress-responsive pathways to improve stress tolerance. Conversely, BnaA10T0057200WE potentially acts as a negative regulator, suppressing growth-related pathways to facilitate stress adaptation. Additionally, distinct BnaTRM genes exhibited early-response, late-response, or biphasic expression patterns, reflecting the functional diversity of TRM family members in plant multilevel stress adaptation mechanisms. Through targeted CRISPR-Cas9-mediated genome editing of genes, we aim to establish genotype–phenotype correlations by quantifying stress tolerance metrics (electrolyte leakage, ROS scavenging capacity) in engineered rapeseed lines and further investigation of their molecular mechanisms and functional validation. It will help elucidate their precise roles in rapeseed stress resistance and provide novel theoretical foundations for molecular breeding.
3.3. Limitations and Future Directions
While this study establishes foundational insights into the biological functions of TRM genes in polyploid plants, several critical questions warrant further investigation: First, dynamic subcellular studies employing live-cell imaging of GFP-tagged TRM proteins are required to delineate TRM-mediated microtubule reorganization during stress recovery phases. Second, protein interaction analyses through co-immunoprecipitation (Co-IP) experiments are necessary to validate TRM interactions with TON1/PP2A subunits in Brassica napus, coupled with systematic characterization of the TTP (TON1-TRM-PP2A) complex’s dynamic behavior and molecular mechanisms in microtubule remodeling. Third, epigenetic profiling should be conducted to identify histone modification signatures (e.g., H3K4me3/H3K27me3) at TRM gene loci. These advanced investigations will systemically integrate omics predictions with functional validation, thereby refining our understanding of TRM regulatory networks and providing precise molecular targets for crop genetic improvement. However, it is crucial to acknowledge that current findings remain confined to in silico predictions; subsequent functional characterization through CRISPR-based overexpression or knockout mutants will be essential to validate gene-specific roles in stress adaptation.
Future development of an integrated BnaTRM database combining proteomic and single-cell transcriptomic datasets will enable systems-level dissection of TRM-mediated signaling networks. Furthermore, integration with CRISPR-based genome editing technologies will facilitate precise plant functional characterization of TRM genes in stress responses. Identification of these key TRM genes provides directly applicable genetic targets for molecular breeding in Brassica napus, establishing foundational resources for developing stress-resilient varieties. For instance, constitutive promoter-driven overexpression of BnaA02T0162800WE can significantly enhance cold, drought, and salt tolerance in elite cultivars, generating transgenic plants with multistress resistance, while CRISPR-mediated knockout of BnaA10T0057200WE could release its negative regulatory effect, markedly improving plant stress tolerance. The translational application of these TRM genetic engineering strategies holds significant socioeconomic value: cultivation of TRM-modified varieties is projected to reduce global rapeseed yield losses caused by abiotic stresses, thereby enhancing agricultural sustainability and production stability.
4. Materials and Methods
4.1. Identification of the TRM Family
The BLASTp program (e < 1 × 10⁻⁵) was used to search for TRM candidate protein sequences (TAIR 10.1) in the whole-genome protein sequences of Brassica napus (Westar.v0) by using Arabidopsis thaliana TRM family proteins as bait sequences. Then, the candidate protein sequence of TRM in Brassica napus was analyzed, and the sequence containing TRM Motif was reserved as a member of the TRM family. Genes that do not conform to TRM family characteristics are removed and visualized by TBTOOLs v2.310 software. In this study, we used the following database to retrieve TRM sequences: TAIR database (TAIR10, https://www.arabidopsis.org/ (accessed on 5 March 2024)), Multiomics information resources of Brassica napus (BNIR) (https://yanglab.hzau.edu.cn/ (accessed on 5 March 2024)), National Center for Biotechnology Information (NCBI, www.ncbi.nlm.nih.gov (accessed on 1 April 2024)), MEME v5.5.7 (Multiple Expectation maximization for Motif Elicitation) (https://meme-suite.org/meme/tools/meme (accessed on 1 April 2024)).
4.2. Chromosome Location
We drew the chromosome map of BnaTRM family genes with TBTOOLs v2.310 software. The genome annotation file Westar.v0.GFF3 used in it comes from the website of BNIR (https://yanglab.hzau.edu.cn/ (accessed on 3 May 2024)).
4.3. Sequence Alignment and Phylogeny Analysis of BnaTRMs
We used MEGA 11.0.13 software to perform a phylogenetic analysis of the TRM families of Brassica napus and Arabidopsis thaliana. The built-in ClustalW program was used for multiple sequence alignments, and the neighbor-joining (NJ) method with a bootstrap value of 1000 repetitions was used to construct the phylogenetic tree. And the iTOL website (https://itol.embl.de/ (accessed on 25 May 2024)) was used to visualize the phylogenetic tree.
4.4. Synteny Analysis of BnaTRM Genes
To investigate the collinearity of BnaTRM genes with other plant species, we employed the One Step McScanX program in TBtools v2.310 to generate collinear files. Genome data of Brassica napus (Westar.v0), Arabidopsis thaliana (TAIR 10.1), soybean (Glycine max v2.1), rice (Oryza sativa L. 1.0), and maize (Zea mays L. 5.0) came from https://plants.ensembl.org/index.html (accessed on 14 June 2024). The results are visualized and built by TBtools v2.310.
4.5. The Biophysical Properties of BnaTRMs
The relative molecular weight (Mw), amino acid number (AA), and theoretical pI of protein, a member of the TRM family gene in Brassica napus, were calculated by ExPasy (https://www.expasy.org (accessed on 24 July 2024)). The subcellular localization of BnaTRM proteins was predicted with the WoLF PSORT online tool (https://wolfpsort.hgc.jp/ (accessed on 24 July 2024)).
4.6. The Identification of Cis-Regulatory Elements in Promoter Regions
To identify the putative cis-acting elements found in BnaTRM genes, the 2.0 kb genomic sequence upstream of the initiation codon (ATG) of each gene was subjected to PlantCARE (https://bioinformatics.psb.ugent.be/webtools/plantcare/html/ (accessed on 28 December 2024)). The results were visualized and constructed using TBtools v2.310.
4.7. RNA Isolation and Quantitative Real-Time PCR (qRT-PCR) Analysis
For gene expression analysis, 7-day-old wild-type Westar seedlings were grown in black plastic culture containers with Hoagland nutrient solution. After 14 days, the seedling roots were exposed to simulated drought stress with 20% PEG-6000 and salt stress with 200 mM NaCl solution. Four treatment time points were set: 0 h (control), 3 h, 12 h, and 48 h. Immediately after each treatment, leaf tissue samples were collected for subsequent RNA extraction. Total RNA was extracted using the Plant (Polysaccharide and Polyphenols) RNA Extraction Kit (Easy-DO, Hangzhou, China), and then we verified the quality and measured concentration. For cDNA Synthesis with the FastKing gDNA Dispelling RT SuperMix (TIANGEN, Beijing, China), qRT-PCR was performed using the UltraSYBR Mixture (Cwbio, Jiangsu, China) with CFX Opus 96 real-time PCR system (BIO-RAD, Hercules, CA, USA). The gene of BnaC09T0180700WE served as a reference gene. Three biological replicates were included in the expression analysis. The relative expression levels of BnaTRMs were calculated using the 2−∆∆Ct approach. Primers are listed in Table S1.
5. Conclusions
This study presents a comprehensive investigation of the TRM gene family in Brassica napus, employing an integrative, multidisciplinary framework that combines bioinformatics, comparative genetics, and rigorous experimental validation to systematically explore the genomic architecture, functional diversity, and agronomic relevance of BnaTRM genes. Regarding the genomic organization, this study reveals that BnaTRM genes exhibit a distinct and uneven chromosomal distribution. Specifically, 87% of these genes are localized to the nucleus, while the remaining 13% are found in chloroplasts or cytoplasm. Furthermore, the genes cluster on chromosomes A09 or C09, suggesting a potential hot spot for TRM gene activity. The extensive synteny observed between BnaTRM genes and those in soybeans underscores the evolutionary conservation of this gene family across species.
The functional diversity of BnaTRM genes is also illuminated in this study. By analyzing promoter-enriched cis-elements, such as ABRE and ARE, we discovered that these elements play a crucial role in regulating gene expression. Additionally, the biphasic expression patterns observed under stress conditions implicate BnaTRM genes in hormone signaling and abiotic stress adaptation. This finding suggests that these genes may play a pivotal role in the plant’s response to environmental stressors. Through integrative analysis of the BnaTRM gene family, this study identified six stress-responsive candidate genes (BnaA02T0162800WE, BnaA10T0057200WE, BnaA03T0002600WE, BnaA05T0045700WE, BnaC04T0062800WE, and BnaA02T0207500WE) as key regulators of abiotic stress adaptation. Based on the results of this paper and previous studies [11,24,29,30], the present study proposes that BnaTRMs form synergistic regulatory modules by interacting with other stress-related proteins (such as MYB family transcription factors, ion transporters, and other related proteins) and are assembled into multisubunit complexes within plant cells. These complexes may dynamically regulate the expression of key genes (e.g., the MYB transcription factor family) in cold, drought, and salt stress response networks, thereby enhancing plant adaptability to multiple environmental stresses.
Collectively, this study advances our understanding of microtubule-mediated stress responses in polyploid crops and establishes a solid foundation for precision breeding strategies aimed at enhancing agricultural sustainability. The insights gained from this research have the potential to revolutionize crop breeding and improve the resilience of Brassica napus and other polyploid crops to environmental stressors, ultimately contributing to global food security and sustainability.
Supplementary Materials
The following supporting information can be downloaded at: https://www.mdpi.com/article/10.3390/plants14121858/s1, Table S1: Primer sequences used for RT-qPCR.
Author Contributions
Y.Z. (Yunlu Zhang), R.W., X.Y., C.Q. and P.Z. designed the research; Y.Z. (Yunlu Zhang), K.Z., H.Z. and J.Z. performed the experiments; Y.Z. (Yunlu Zhang), K.Z. and C.Q. analyzed data; Y.Z. (Yunlu Zhang), K.Z., Y.Z. (Yang Zhu) and C.Q. wrote the manuscript. All authors have read and agreed to the published version of the manuscript.
Funding
This research was supported by the Hangzhou Joint Fund of the Zhejiang Provincial Natural Science Foundation of China under Grant No. LHZY24C140001.
Data Availability Statement
The data presented in this study are available in the paper and Supplementary Files.
Conflicts of Interest
The authors declare no conflicts of interest.
References
- Blackshaw, R.E.; Johnson, E.N.; Gan, Y.; May, W.E.; McAndrews, D.W.; Barthet, V.; McDonald, T.; Wispinski, D. Alternative oilseed crops for biodiesel feedstock on the Canadian prairies. Can. J. Plant Sci. 2011, 91, 889–896. [Google Scholar] [CrossRef] [Scilit]
- Bayer, P.E.; Hurgobin, B.; Golicz, A.A.; Chan, C.-K.K.; Yuan, Y.; Lee, H.; Renton, M.; Meng, J.; Li, R.; Long, Y.; et al. Assembly and comparison of two closely related Brassica napus genomes. Plant Biotechnol. J. 2017, 15, 1602–1610. [Google Scholar] [CrossRef] [Scilit]
- Chalhoub, B. Early allopolyploid evolution in the post-Neolithic Brassica napus oilseed genome. Science 2014, 345, 1255, Erratum in Science 2014, 348, 1260782. [Google Scholar] [CrossRef] [Scilit]
- Sun, F.; Fan, G.; Hu, Q.; Zhou, Y.; Guan, M.; Tong, C.; Li, J.; Du, D.; Qi, C.; Jiang, L.; et al. The high-quality genome of Brassica napus cultivar ‘ZS11’ reveals the introgression history in semi-winter morphotype. Plant J. 2017, 92, 452–468. [Google Scholar] [CrossRef] [Scilit]
- Drevensek, S.; Goussot, M.; Duroc, Y.; Christodoulidou, A.; Steyaert, S.; Schaefer, E.; Duvernois, E.; Grandjean, O.; Vantard, M.; Bouchez, D.; et al. The Arabidopsis TRM1-TON1 Interaction Reveals a Recruitment Network Common to Plant Cortical Microtubule Arrays and Eukaryotic Centrosomes. Plant Cell 2012, 24, 178–191. [Google Scholar] [CrossRef] [Scilit]
- Camilleri, C.; Azimzadeh, J.; Pastuglia, M.; Bellini, C.; Grandjean, O.; Bouchez, D. The Arabidopsis TONNEAU2 gene encodes a putative novel protein phosphatase 2A regulatory subunit essential for the control of the cortical cytoskeleton. Plant Cell 2002, 14, 833–845. [Google Scholar] [CrossRef] [Scilit]
- Ehrhardt, D.W.; Shaw, S.L. Microtubule dynamics and organization in the plant cortical array. Annu. Rev. Plant Biol. 2006, 57, 859–875. [Google Scholar] [CrossRef] [Scilit]
- Duroc, Y.; Bouchez, D.; Pastuglia, M. The Preprophase Band and Division Site Determination in Land Plants. In The Plant Cytoskeleton; Liu, B., Ed.; Springer: New York, NY, USA, 2011; pp. 145–185. [Google Scholar]
- Landrein, B.; Hamant, O. How mechanical stress controls microtubule behavior and morphogenesis in plants: History, experiments and revisited theories. Plant J. 2013, 75, 324–338. [Google Scholar] [CrossRef] [Scilit]
- Wade, R.H. On and Around Microtubules: An Overview. Mol. Biotechnol. 2009, 43, 177–191. [Google Scholar] [CrossRef] [Scilit]
- Azimzadeh, J.; Nacry, P.; Christodoulidou, A.; Drevensek, S.; Camilleri, C.; Amiour, N.; Parcy, F.; Pastuglia, M.; Bouchez, D. Arabidopsis TONNEAU1 proteins are essential for preprophase band formation and interact with centrin. Plant Cell 2008, 20, 2146–2159. [Google Scholar] [CrossRef] [Scilit]
- Lee, Y.K.; Kim, G.-T.; Kim, I.-J.; Park, J.; Kwak, S.-S.; Choi, G.; Chung, W.-I. LONGIFOLIA1 and LONGIFOLIA2, two homologous genes, regulate longitudinal cell elongation in Arabidopsis. Development 2006, 133, 4305–4314. [Google Scholar] [CrossRef] [Scilit]
- Schmitz, A.J.; Begcy, K.; Sarath, G.; Walia, H. Rice Ovate Family Protein 2 (OFP2) alters hormonal homeostasis and vasculature development. Plant Sci. 2015, 241, 177–188. [Google Scholar] [CrossRef] [Scilit]
- Zhan, P.; Ma, S.; Xiao, Z.; Li, F.; Wei, X.; Lin, S.; Wang, X.; Ji, Z.; Fu, Y.; Pan, J.; et al. Natural variations in grain length 10 (GL10) regulate rice grain size. J. Genet. Genomics 2022, 49, 405–413. [Google Scholar] [CrossRef] [Scilit]
- Wu, S.; Zhang, B.; Keyhaninejad, N.; Rodriguez, G.R.; Kim, H.J.; Chakrabarti, M.; Illa-Berenguer, E.; Taitano, N.K.; Gonzalo, M.J.; Diaz, A.; et al. A common genetic mechanism underlies morphological diversity in fruits and other plant organs. Nat. Commun. 2018, 9, 4734. [Google Scholar] [CrossRef] [Scilit]
- Bao, Z.; Guo, Y.; Deng, Y.; Zang, J.; Zhang, J.; Deng, Y.; Ouyang, B.; Qu, X.; Buerstenbinder, K.; Wang, P. Microtubule-associated protein SlMAP70 interacts with IQ67-domain protein SlIQD21a to regulate fruit shape in tomato. Plant Cell 2023, 35, 4266–4283. [Google Scholar] [CrossRef] [Scilit]
- Bao, Z.; Xu, Z.; Zang, J.; Buerstenbinder, K.; Wang, P. The Morphological Diversity of Plant Organs: Manipulating the Organization of Microtubules May Do the Trick. Front. Cell Dev. Biol. 2021, 9, 649626. [Google Scholar] [CrossRef] [Scilit]
- McNally, F.J.; Roll-Mecak, A. Microtubule-severing enzymes: From cellular functions to molecular mechanism. J. Cell Biol. 2018, 217, 4057–4069. [Google Scholar] [CrossRef] [Scilit]
- Schaefer, E.; Belcram, K.; Uyttewaal, M.; Duroc, Y.; Goussot, M.; Legland, D.; Laruelle, E.; de Tauzia-Moreau, M.-L.; Pastuglia, M.; Bouchez, D. The preprophase band of microtubules controls the robustness of division orientation in plants. Science 2017, 356, 186–189. [Google Scholar] [CrossRef] [Scilit]
- Spinner, L.; Gadeyne, A.; Belcram, K.; Goussot, M.; Moison, M.; Duroc, Y.; Eeckhout, D.; De Winne, N.; Schaefer, E.; Van de Slijke, E.; et al. A protein phosphatase 2A complex spatially controls plant cell division. Nat. Commun. 2013, 4, 1863. [Google Scholar] [CrossRef] [Scilit]
- Guo, Q.; Ng, P.Q.; Shi, S.; Fan, D.; Li, J.; Zhao, J.; Wang, H.; David, R.; Mittal, P.; Trung, D.; et al. Arabidopsis TRM5 encodes a nuclear-localised bifunctional tRNA guanine and inosine-N1-methyltransferase that is important for growth. PLoS ONE 2019, 14, e0225064. [Google Scholar] [CrossRef] [Scilit]
- Tang, J.; Jia, P.; Xin, P.; Chu, J.; Shi, D.-Q.; Yang, W.-C. The Arabidopsis TRM61/TRM6 complex is a bona fide tRNA N1-methyladenosine methyltransferase. J. Exp. Bot. 2020, 71, 3024–3036. [Google Scholar] [CrossRef] [Scilit]
- Wu, L.; Chen, X.; Zhang, P.; Yan, S.; Zhang, T.; Li, Y. TON1 recruiting motif 21 positively regulates the flavonoid metabolic pathway at the translational level in Arabidopsis thaliana. Planta 2024, 259, 65. [Google Scholar] [CrossRef] [Scilit]
- Wang, S.; Li, S.; Liu, Q.; Wu, K.; Zhang, J.; Wang, S.; Wang, Y.; Chen, X.; Zhang, Y.; Gao, C.; et al. The OsSPL16-GW7 regulatory module determines grain shape and simultaneously improves rice yield and grain quality. Nat. Genet. 2015, 47, 949–954. [Google Scholar] [CrossRef] [Scilit]
- Wang, Y.; Xiong, G.; Hu, J.; Jiang, L.; Yu, H.; Xu, J.; Fang, Y.; Zeng, L.; Xu, E.; Xu, J.; et al. Copy number variation at the GL7 locus contributes to grain size diversity in rice. Nat. Genet. 2015, 47, 944–948. [Google Scholar] [CrossRef] [Scilit]
- Liu, J.; Van Eck, J.; Cong, B.; Tanksley, S.D. A new class of regulatory genes underlying the cause of pear-shaped tomato fruit. Proc. Natl. Acad. Sci. USA 2002, 99, 13302–13306. [Google Scholar] [CrossRef] [Scilit]
- Van Der Knaap, E.; Lippman, Z.B.; Tanksley, S.D. Extremely elongated tomato fruit controlled by four quantitative trait loci with epistatic interactions. TAG Theor. Appl. Genet. Theor. Angew. Genet. 2002, 104, 241–247. [Google Scholar] [CrossRef] [Scilit]
- Rodriguez, G.R.; Kim, H.J.; van der Knaap, E. Mapping of two suppressors of OVATE (sov) loci in tomato. Heredity 2013, 111, 256–264. [Google Scholar] [CrossRef] [Scilit]
- Zhang, B.; Li, Q.; Keyhaninejad, N.; Taitano, N.; Sapkota, M.; Snouffer, A.; van der Knaap, E. A combinatorial TRM-OFP module bilaterally fine-tunes tomato fruit shape. New Phytol. 2023, 238, 2393–2409. [Google Scholar] [CrossRef] [Scilit]
- Zhao, L.; Wang, K.; Wang, Z.; Chu, S.; Chen, C.; Wang, L.; Ren, Z. Pan-Genome Analysis of TRM Gene Family and Their Expression Pattern under Abiotic and Biotic Stresses in Cucumber. Horticulturae 2024, 10, 908. [Google Scholar] [CrossRef] [Scilit]
- Shokri-Gharelo, R.; Noparvar, P.M. Molecular response of canola to salt stress: Insights on tolerance mechanisms. PeerJ 2018, 6, e4822. [Google Scholar] [CrossRef] [Scilit]
- Ma, H.; Liu, M. The microtubule cytoskeleton acts as a sensor for stress response signaling in plants. Mol. Biol. Rep. 2019, 46, 5603–5608. [Google Scholar] [CrossRef] [Scilit]
- Kerr, G.P.; Carter, J.V. Relationship between Freezing Tolerance of Root-Tip Cells and Cold Stability of Microtubules in Rye (Secale cereale L. cv Puma). Plant Physiol. 1990, 93, 77–82. [Google Scholar] [CrossRef] [Scilit]
- Wang, C.; Li, J.; Yuan, M. Salt tolerance requires cortical microtubule reorganization in Arabidopsis. Plant Cell Physiol. 2007, 48, 1534–1547. [Google Scholar] [CrossRef] [Scilit]
- Zhang, Y.; Yang, Z.; He, Y.; Liu, D.; Liu, Y.; Liang, C.; Xie, M.; Jia, Y.; Ke, Q.; Zhou, Y.; et al. Structural variation reshapes population gene expression and trait variation in 2105 Brassica napus accessions. Nat. Genet. 2024, 56, 2538–2550. [Google Scholar] [CrossRef] [Scilit]
- Ding, Y.; Feng, L.; Li, P.; Yang, X.; Li, M.; Liu, H.; Xu, J.; Zhang, J.; Sun, S.; Zhou, X.; et al. The Alpha/Beta-Hydrolase Fold Superfamily in Brassica napus: Expression Profiles and Functional Implications of Clade-3 BnABH Proteins in Response to Abiotic Stress. Int. J. Mol. Sci. 2025, 26, 4746. [Google Scholar] [CrossRef] [Scilit]
- Wang, R.; Wu, G.; Zhang, J.; Hu, W.; Yao, X.; Jiang, L.; Zhu, Y. Integration of GWAS and transcriptome analysis to identify temperature-dependent genes involved in germination of rapeseed (Brassica napus L.). Front. Plant Sci. 2025, 16, 1551317. [Google Scholar] [CrossRef] [Scilit]
- Wu, G.; Zhou, Y.; Zhang, J.; Gong, M.; Jiang, L.; Zhu, Y. Genome-wide association study and candidate gene identification for the cold tolerance at the seedling stage of rapeseed (Brassica napus L.). Crop Des. 2025, 4, 100083. [Google Scholar] [CrossRef] [Scilit]
- Gu, Y.; Zhao, Q.-C.; Sun, D.-L.; Song, W.-Q. Construction of genetic linkage map and localization of NBS-LRR like resistance gene analogues in cauliflower (Brassica oleracea var. botrytis). Yi Chuan = Hereditas 2007, 29, 751–757. [Google Scholar] [CrossRef] [Scilit]
- Liu, M.; Chang, W.; Fan, Y.; Sun, W.; Qu, C.; Zhang, K.; Liu, L.; Xu, X.; Tang, Z.; Li, J.; et al. Genome-Wide Identification and Characterization of NODULE-INCEPTION-Like Protein (NLP) Family Genes in Brassica napus. Int. J. Mol. Sci. 2018, 19, 2270. [Google Scholar] [CrossRef] [Scilit]
- Song, X.; Lu, X.; Shen, Y.; Guo, S.; Guan, Y. A modified supercritical Dubinin-Radushkevich model for the accurate estimation of high pressure methane adsorption on shales. Int. J. Coal Geol. 2018, 193, 1–15. [Google Scholar] [CrossRef] [Scilit]
- Wang, T.; Hu, J.; Ma, X.; Li, C.; Yang, Q.; Feng, S.; Li, M.; Li, N.; Song, X. Identification, evolution and expression analyses of whole genome-wide TLP gene family in Brassica napus. BMC Genomics 2020, 21, 264. [Google Scholar] [CrossRef] [Scilit] [PubMed]

Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2025 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license (https://creativecommons.org/licenses/by/4.0/).