


Abstract
We reported copper-catalyzed the 1,4- or 1,6-conjugate additions of arylidenemalonitriles or para-quinone methides with 2-bromodifluoromethylbenzo-1,3-thiazole for the preparation of benzothiazolyldifluoroalkyl containing (di)arylmethine compounds in moderate to excellent yields, providing a facile route for the synthesis of gem-difluoroalkylatated (di)arylmethine compounds. The synthetic utility of the current method was also demonstrated by a gram-scale reaction.
1. Introduction
The selective introduction of a fluoroalkyl group into biologically active molecules has emerged as a reliable strategy to modulate pharmacological properties in drug discovery and development [1,2,3,4,5,6]. Therefore, establishing new efficient methods for the synthesis of fluoroalkyl-containing compounds is currently a hot topic in the field of synthetic chemistry. The incorporation of a gem-difluoroalkyl group substantially influences the electronic properties of its neighboring groups, which may produce beneficial effects such as an enhancement in lipophilicity, metabolic stability, bioavailability [7]. For instance, eflornithine [8] and gemcitabine [9] are drugs containing a gem-difluoroalkyl group. Furthermore, the difluoromethylene analogs of both Vitamin D [10] and docetaxel [11] show improved properties. Despite considerable progress in selective trifluoromethylation [12], the development of methods for selective gem-difluoroalkylation is still highly desirable [13,14,15,16,17].
2-Bromodifluoromethylbenzo-1,3-thiazoles 1, as stable and readily available difluoroalkyl reagents for the selective introduction of gem-difluoromethylene benzo-1,3-thiazole moiety, have gained increasing attention in the past decade. A variety of reactions using 1 as a substrate have been reported, including the SRN1 reaction [18,19], aldol reaction [20], cross-coupling reaction [21,22,23,24], (di)functionalization of alkenes reaction [25,26,27,28,29,30], and Truce–Smiles rearrangement reaction [31].
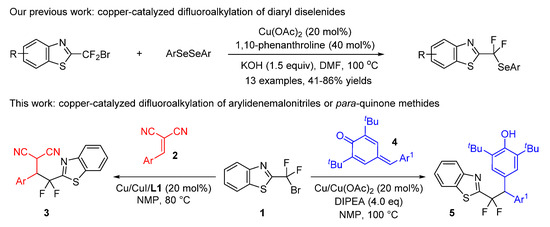
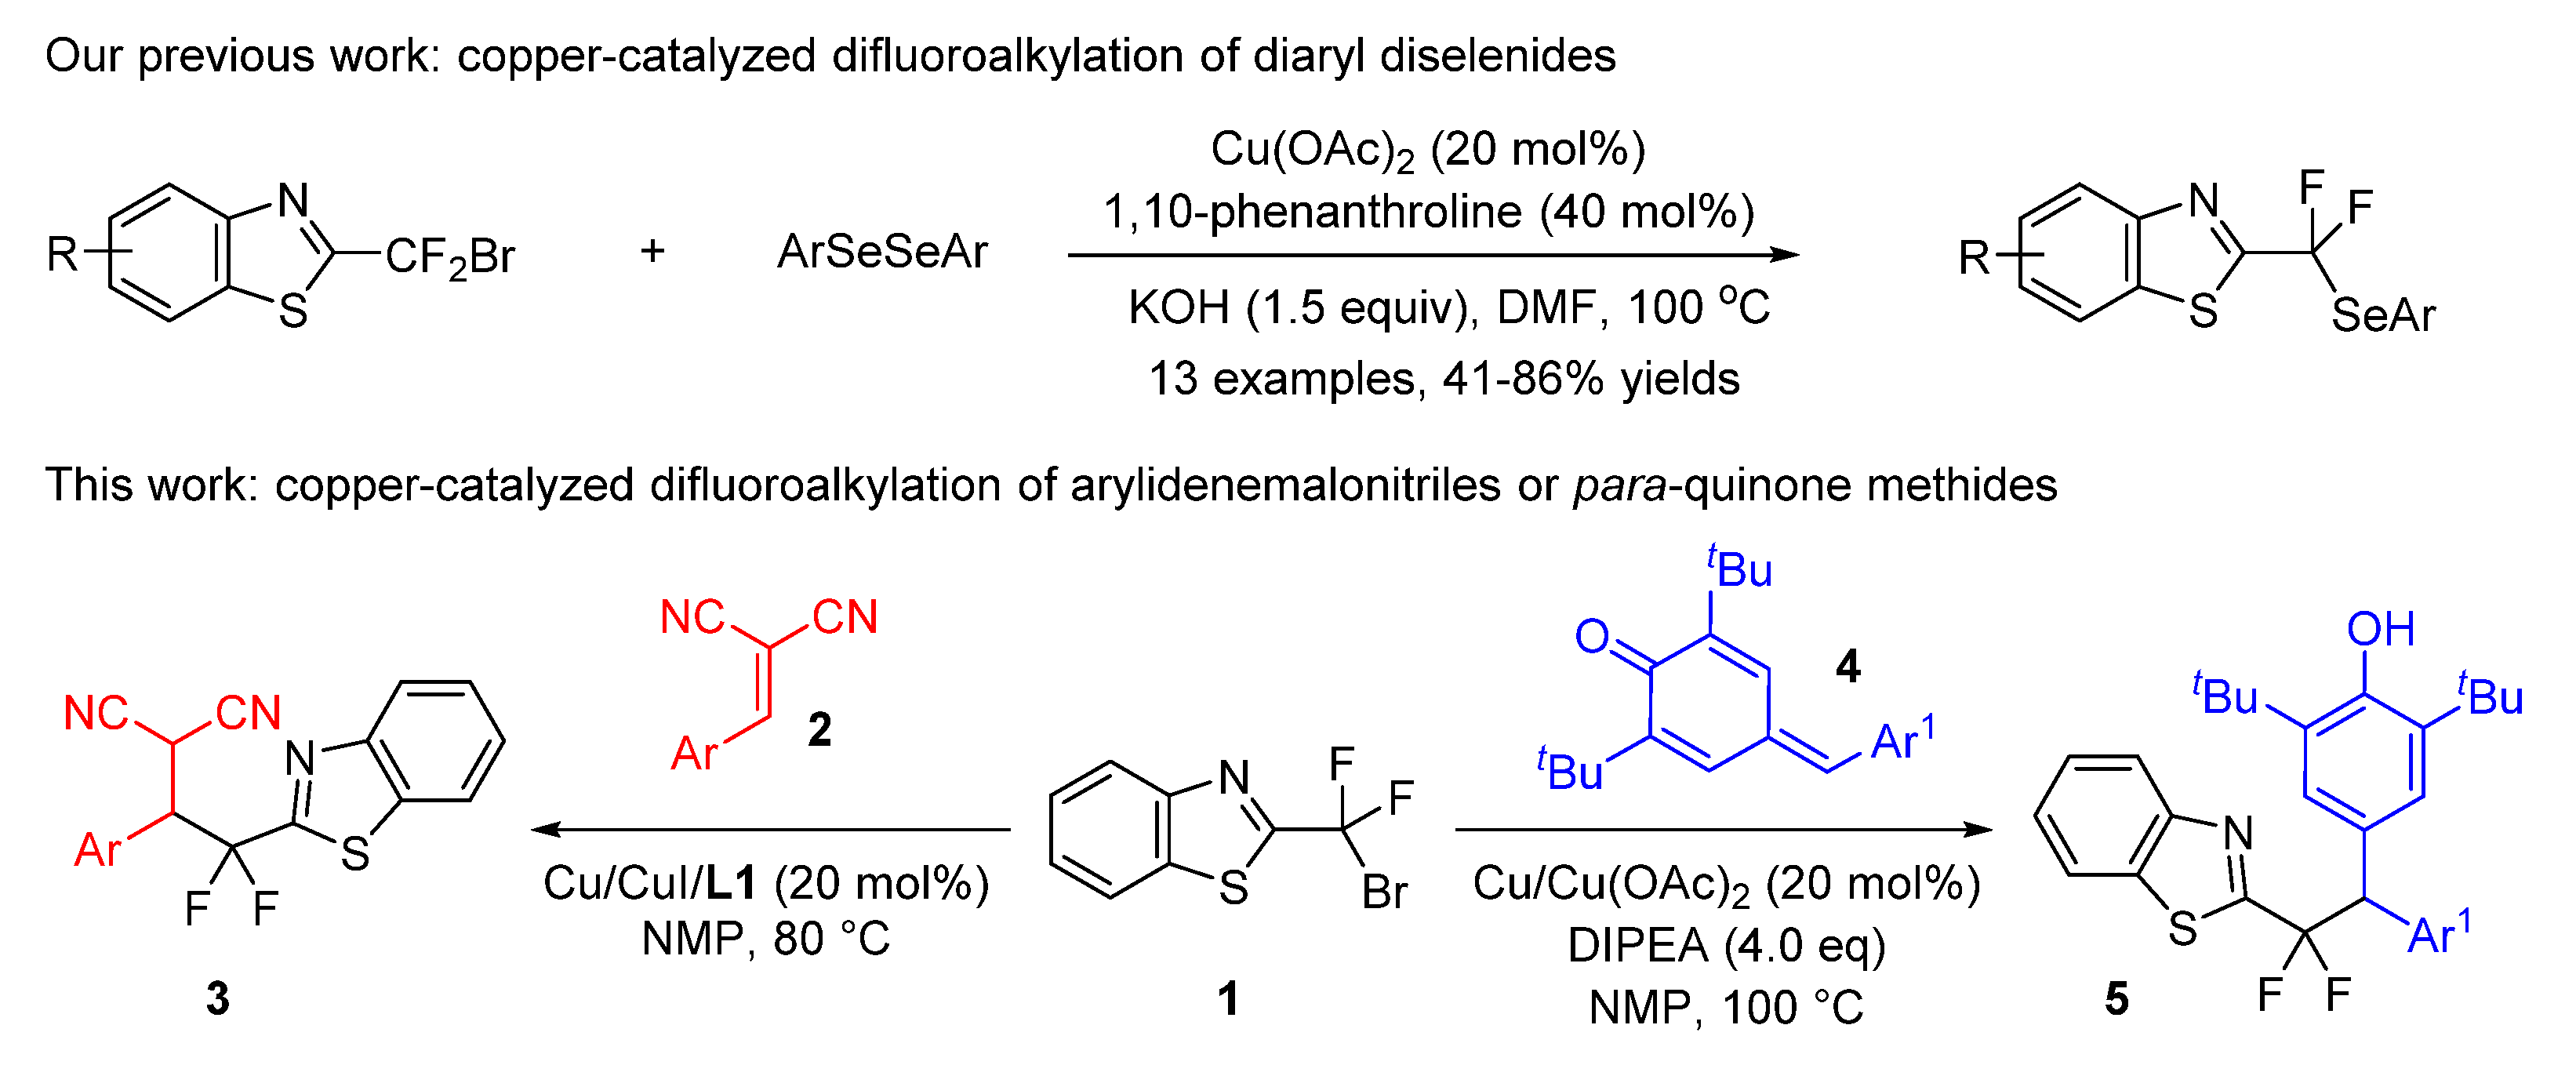
The significance of functional malononitrile [32,33] and diarylmethine frameworks scaffolds [34,35,36,37] has been studied, while combining the gem-difluoroalkyl group with arylidenemalonitriles or para-quinone methides (p-QMs) scaffold remains less explored [38,39,40,41,42]. Given the importance of difluoroalkylated (di)arylmethine skeletons in medicinal research, it is highly important to combine them in one molecule for future drug development. As a continuation of our interest in selective fluoroalkylation with 2-Bromodifluoromethylbenzo-1,3-thiazoles 1 [43,44], we have recently completed the efficient synthesis of 1 and its aryl selenylation to furnish gem-difluoromethyl linked aryl seleno compounds [45], as shown in Scheme 1. Herein, the copper-catalyzed 1,4 or 1,6-conjugate addition reaction of 1 with arylidenemalonitriles or p-QMs was developed to furnish gem-difluoroalkylated (di)arylmethine compounds.
Scheme 1.
The synthetic application of 2-bromodifluoromethylbenzo-1,3-thiazole.
2. Results
2.1. The Reaction Conditions Optimization of 1 and 2
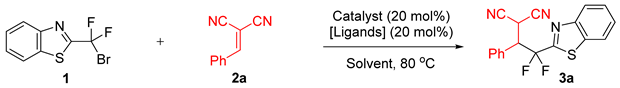
Initially, we chose 2-Bromodifluoromethylbenzo-1,3-thiazole 1 and 2-benzylidenemalononitrile 2a as the model substrates to investigate reaction conditions. As outlined in Table 1, firstly, using 20 mol% CuI as the catalyst only, the product 3a was obtained in only an 18% yield (Table 1, entry 1).
Table 1.
Optimization reaction conditions of 1 and 2 a.
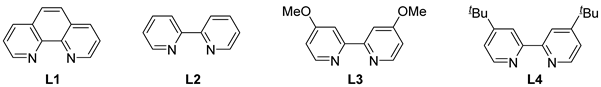
Then, we added four common N,N-bidentate ligands to the reaction and found 1,10-phenanthroline (L1) better than other dipyridyl ligands (39% yield, Table 1, entry 2 vs. 3–5). Using CuI/L1 complex as the catalyst, we then screened different solvents and temperatures. After the screening of various solvents, N-methyl-2-pyrrolidone (NMP) proved to be better than other solvents, such as N,N-dimethylacetamide (DMAc), dimethyl sulfoxide (DMSO), acetonitrile, tetrahydrofuran, and toluene, providing the desired product 3a in a 57% yield (Table 1, entry 8). Whether it was due to raising the reaction temperature to 100 °C or lowering the reaction temperature to 60 °C, the yield of 3a decreased (Table 1, entry 8 vs. 9–10).
A series of copper catalysts [e.g., CuBr, CuCl, and Cu(OAc)2] were also tested (Table 1, entries 11–13), and CuI was determined to be the best. It is worth mentioning that the addition of 20 mol% copper powder was beneficial for improving the yield of 3a from 57% to 81% (Table 1, entry 14 vs. 8). In addition, increasing or decreasing the amount of copper powder both diminished the reaction yield of 3a (Table 1, entry 14 vs. 15–16).
2.2. Reaction Conditions Optimization of 1 and 4
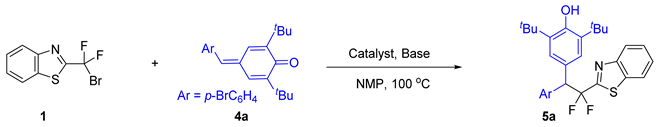
We then investigated the reaction conditions of 2-Bromodifluoromethylbenzo-1,3-thiazole 1 with para-quinone methides (p-QMs) 4 for the synthesis of diarylmethine frameworks, as outlined in Table 2.
Table 2.
Optimization reaction conditions of 1 and 4 a.
Firstly, by using 20 mol% Cu/CuI/L1 as the catalyst, the product 5a was obtained in a 43% yield (Table 2, entry 1). When the amount of copper powder was increased to 1.0 equivalent, the yield of 5a increased to 53% (Table 2, entry 2). On this basis, we investigated different copper salts and ligands and found that Cu(OAc)2 was better (Table 2, entry 4). Further investigation revealed that the ligands were not necessary (Table 2, entry 8 vs. 4).
Next, the addition of bases could increase the yield of 5a, and adding 4.0 equivalents of DIPEA was necessary (Table 2, entry 14). Similarly, both increasing and decreasing the temperature decreased the reaction yield (Table 2, entry 14 vs. 16–17). It was crucial to use both the copper powder and copper salt simultaneously, and the product yield significantly decreased without either (Table 2, entry 14 vs. 18–19). To our surprise, increasing the concentration of NMP could reduce the amount of copper powder to 20 mol% while maintaining almost the same yield of 5a (Table 2, entry 21 vs. 14).
2.3. Substrate Scope
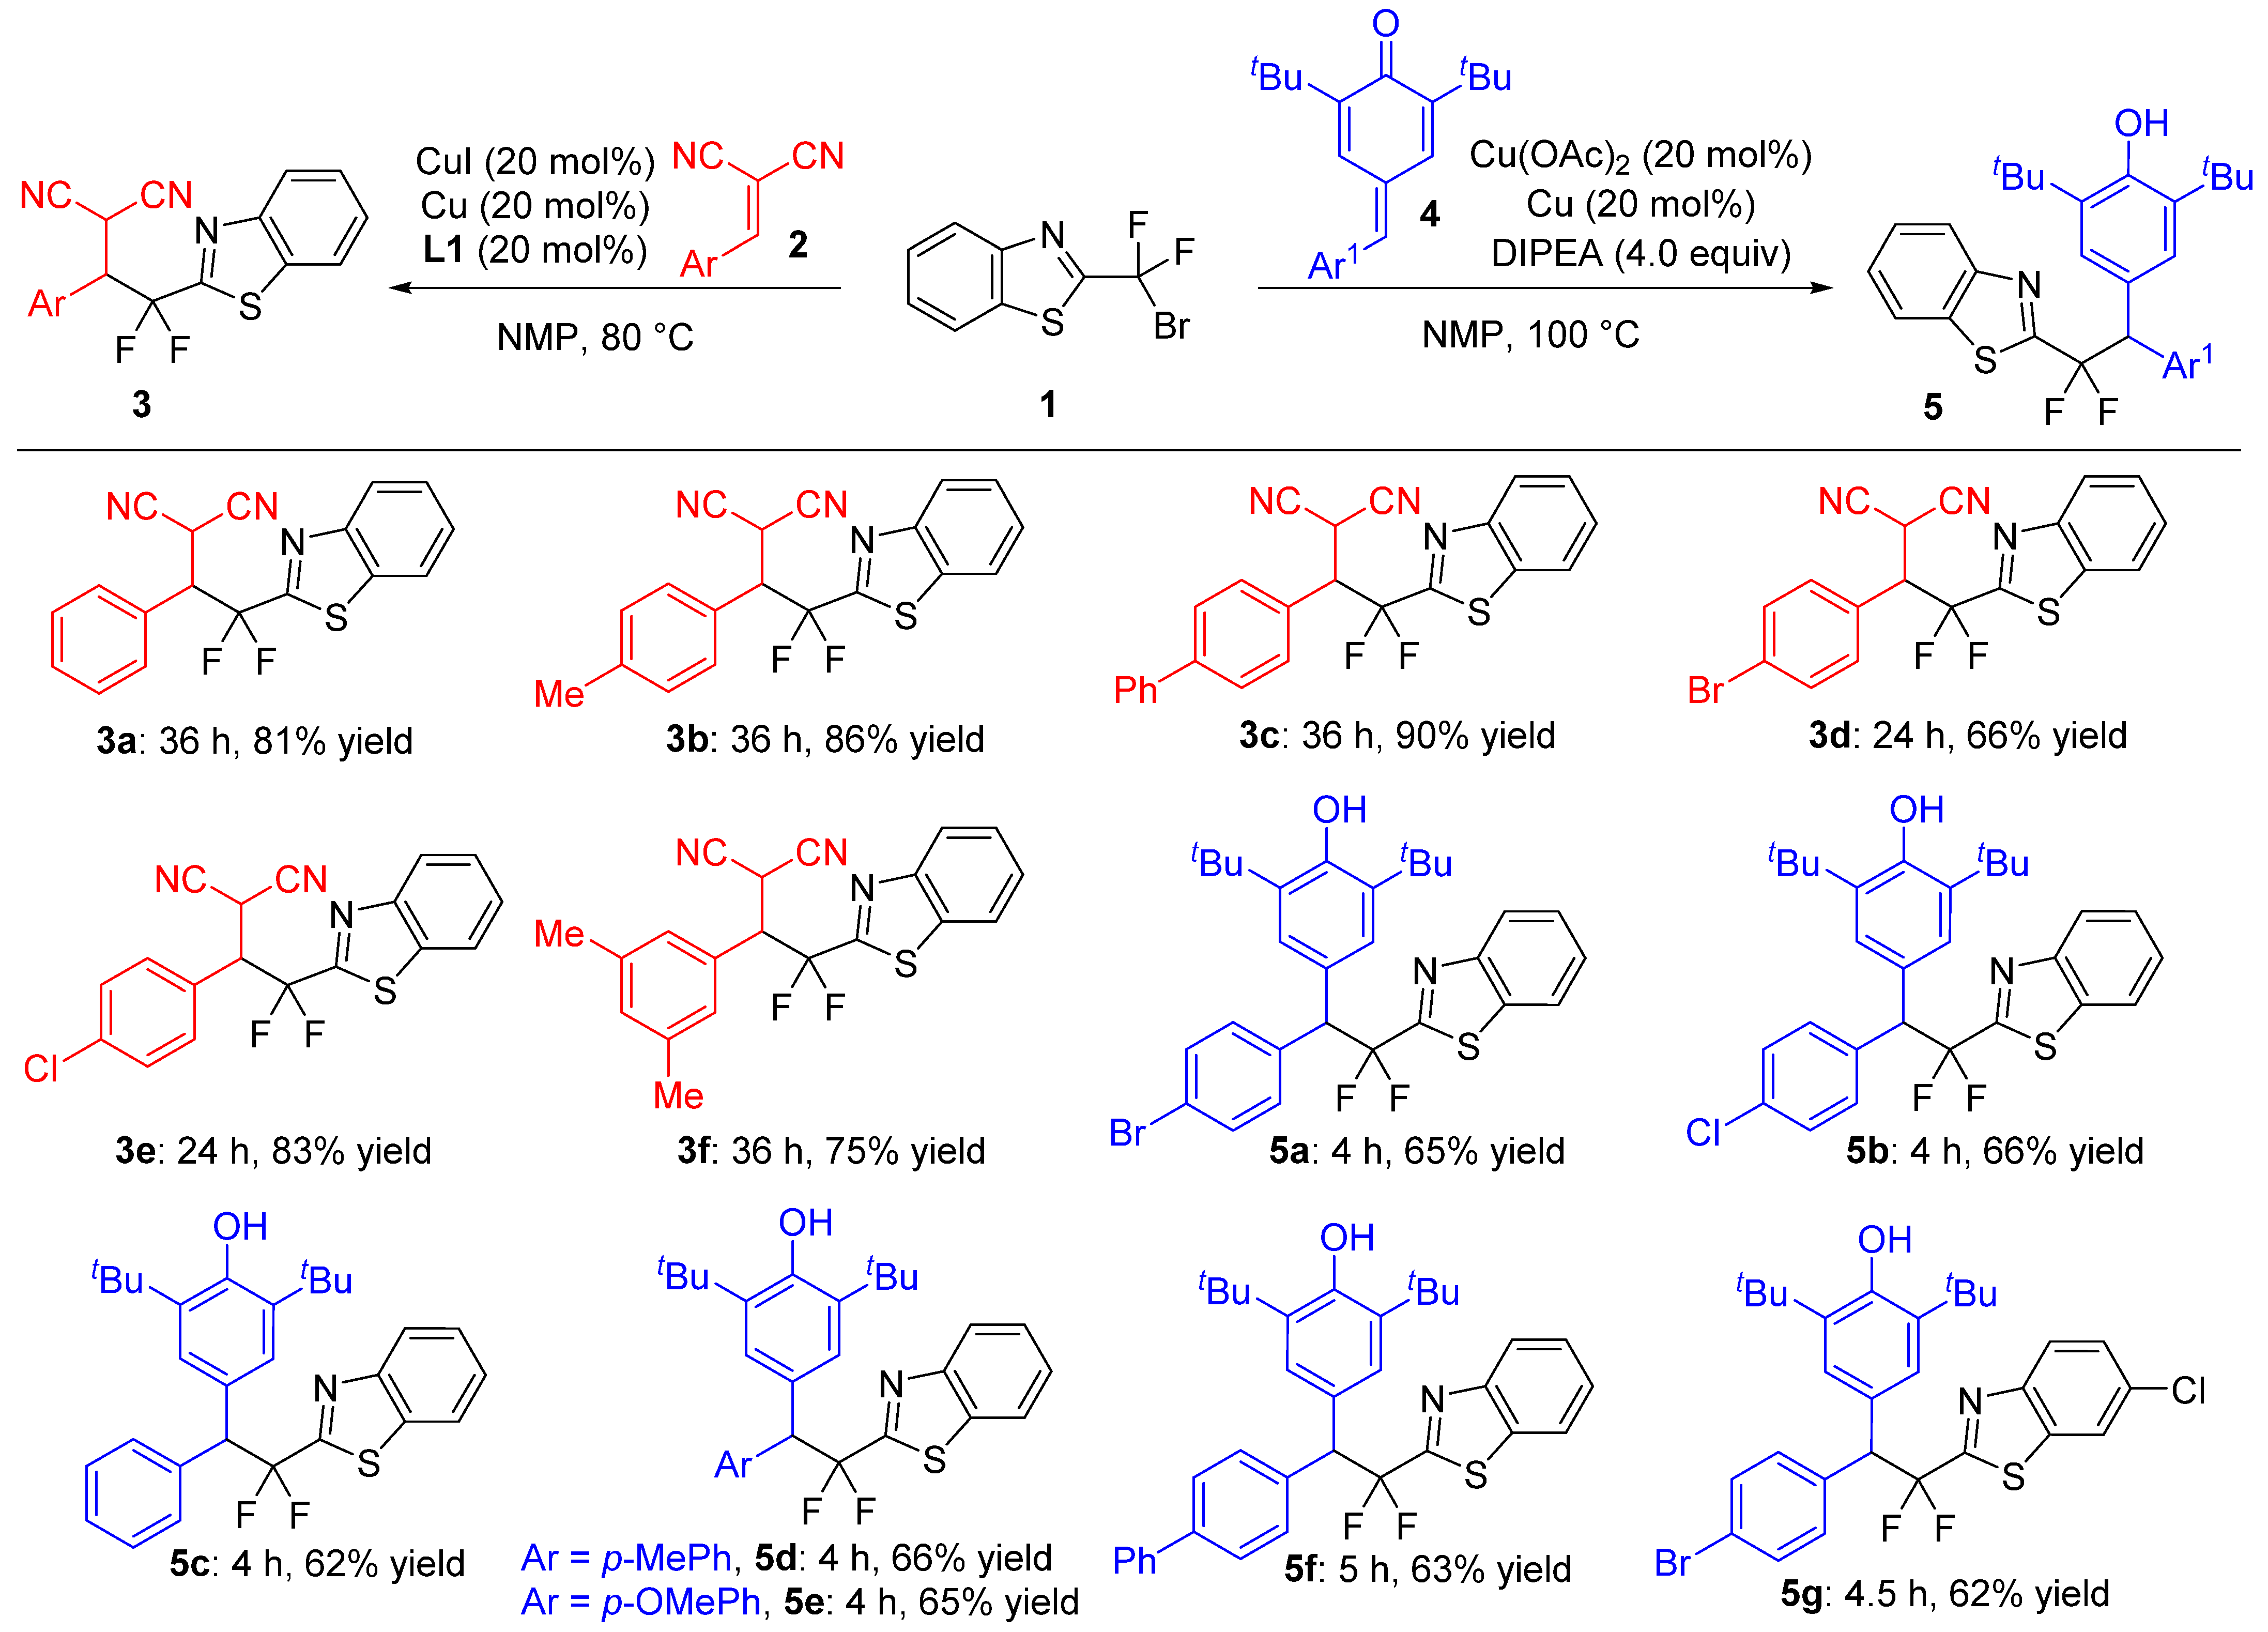
With the optimized conditions in hand, we investigated the generality of this reaction. As shown in Table 3, the substrate scope with respect to arylidenemalonitriles 2 was examined. And, we found that the different nature and position of the substituted arylidenemalonitriles 2 all reacted well with the 2-Bromodifluoromethylbenzo-1,3-thiazole 1 to give rise to the corresponding products 3a–3f in 66–90% yields. We further developed the reaction of 1 with para-quinone methides (p-QMs) 4 for diarylmethine synthesis frameworks. As shown in Table 3, the substrate scope with respect to p-QMs was examined. And, we found that aryl-substituted p-QMs 4 all reacted well with the 2-Bromodifluoromethylbenzo-1,3-thiazole 1 to give rise to the corresponding gem-difluoroalkyl-substituted diarylmethine frameworks 5a–5g in 62–66% yields, whatever the nature of the substituent on the phenyl ring.
Table 3.
Substrate scope for the difluoroalkylation of 2 or 4 with 1.
2.4. Gram-Scale Synthesis
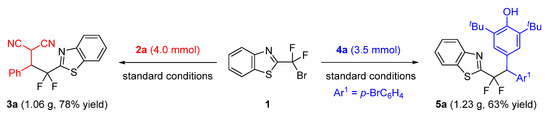
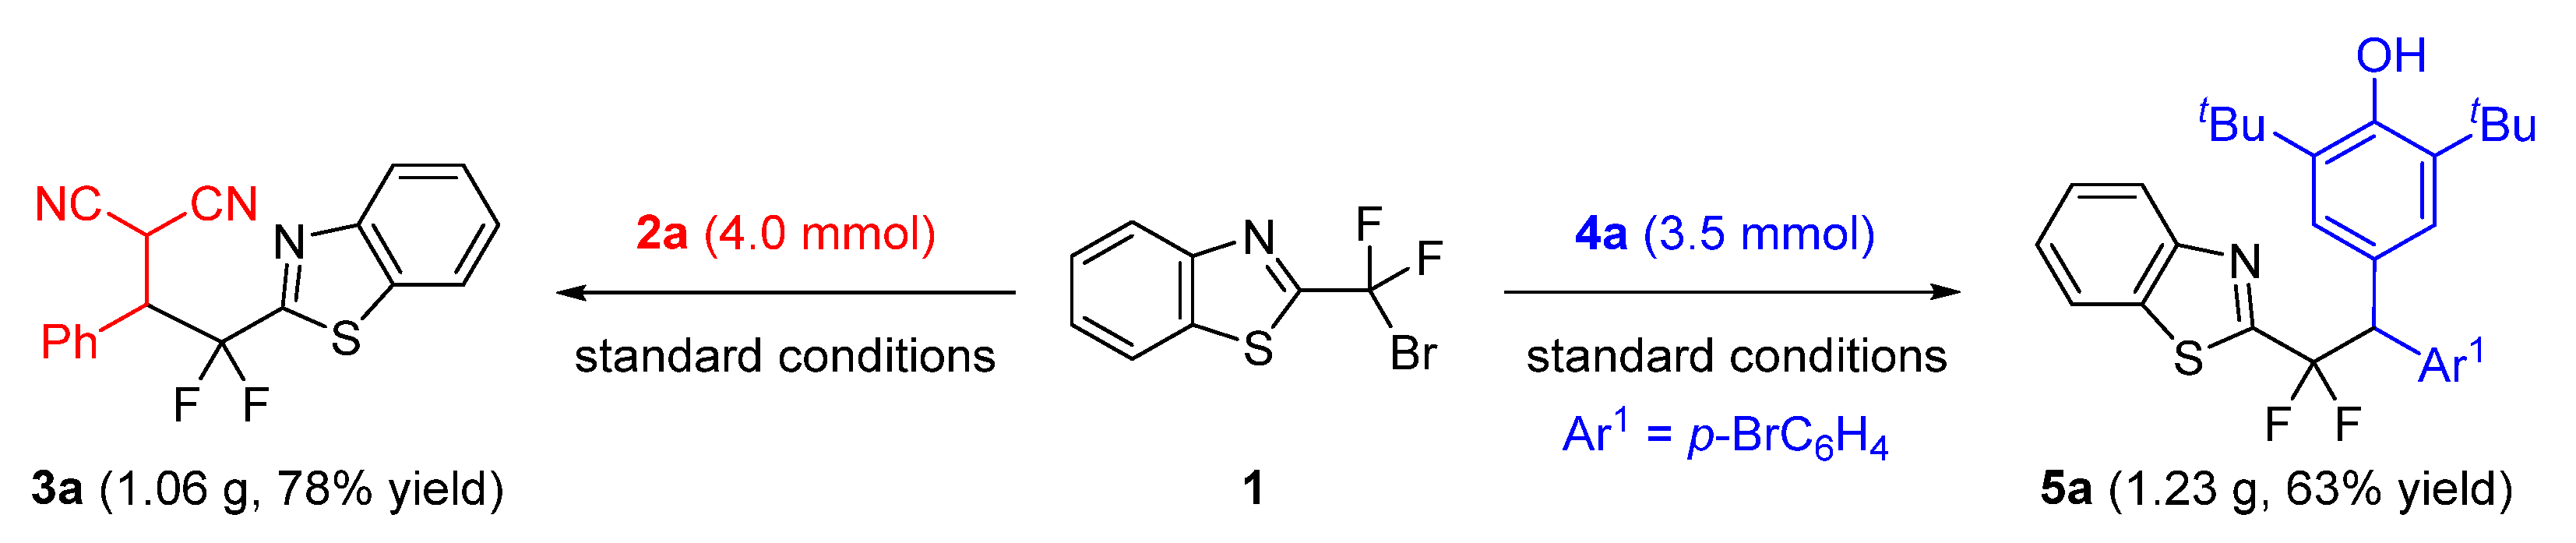
Subsequently, in order to examine the practicality and simplicity of the developed method, the gram-scale reaction was conducted, as Shown in Scheme 2. To our delight, the desired product 3a was obtained in a 78% yield (1.06 g) and the 5a was furnished in a 63% yield (1.23 g).
Scheme 2.
Gram-scale synthesis.
2.5. The Proposed Mechanism
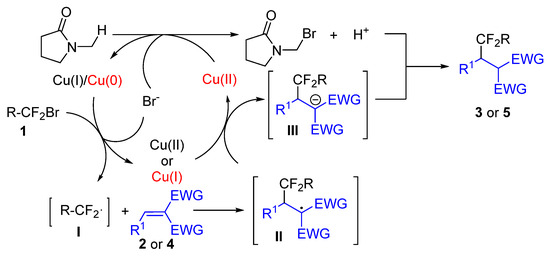
It has previously been shown that a copper catalyst can initiate the generation of radicals from difluoroalkyl halides through single electron transfer. By employing 2.0 eq of TEMPO as the radical scavenger under standard reaction conditions, the products 3a and 5a were not obtained. This result suggests that free radical pathways may be involved in this reaction. We tried to obtain some information about active intermediates. Unfortunately, this idea could not be confirmed.
Based on the above experimental data and the mechanisms reported in the literature, the proposed reaction mechanisms are described in Scheme 3. Firstly, a copper catalyst can initiate the generation of difluoroalkyl radicals I from 1 through single electron transfer [46,47,48,49] upon the loss of bromine, which undergoes an additional reaction with the active olefins 2 or 4 to form intermediate II. Secondly, II is then reduced by copper(I) to form intermediate III and copper(II). Lastly, the copper(II) is reduced by the solvent NMP to the copper catalyst, along with the generation of protons, which then combine with intermediate III to generate product 3 or 5. In the reaction of 1 with 4, on the one hand, Cu(II) may be reduced by copper powder to generate Cu(I), and, on the other hand, it may also serve as a Lewis acid activation for the substrate 4.
Scheme 3.
Proposed mechanism for the copper-catalyzed difluoroalkylation.
3. Materials and Methods
All the commercially available reagents were purchased from Shanghai Energy Chemical and used without further purification. The solvents were treated prior to use according to standard methods. All the reactions were performed under nitrogen using solvents dried by standard methods. NMR spectra were recorded on a Bruker Avance AV400 (400/100/376 MHz 1H/13C/19F) spectrometer (Bruker, Billerica, MA, USA). The 1H and 13C chemical shifts were reported in ppm relative to the residual solvent peak. HR- MS data were obtained using a Thermo Scientific Q Exactive Orbitrap Mass Spectrometer. Silica gel (200–300 mesh) was used for the chromatographic separations.
General Procedure A: All the catalytic reactions were performed in a 10 mL Schlenk tube under N2. Typically, a mixture of 2-Bromodifluoromethyl-benzo-1,3-thiazole 1 (0.45 mmol), arylidenemalonitriles 2 (0.30 mmol), CuI (0.06 mmol), Cu (0.06 mmol), and 1,10-Phenanthroline (L1, 0.06 mmol) was used, with the following addition of 3.0 mL of anhydrous NMP as a solvent. The reaction mixture was stirred at 80 °C until the full disappearance of 1 by TLC analysis (24–36 h), and, then, it was purified by flash column chromatography (petroleum ether:ethyl acetate = 7:1), affording the corresponding product 3.
General Procedure B: All the catalytic reactions were performed in a 10 mL Schlenk tube under N2. Typically, a mixture of 2-Bromodifluoromethyl-benzo-1,3-thiazole 1 (0.20 mmol), p-QMS 4 (0.20 mmol), Cu(OAc)2 (0.04 mmol), Cu (0.04 mmol), and DIPEA (0.80 mmol) was used, with the following addition of 1.0 mL of anhydrous NMP as a solvent. The reaction mixture was stirred at 100 °C until the full disappearance of 1 by TLC analysis (4–5 h), and, then, it was purified by flash column chromatography (petroleum ether:ethyl acetate = 100:1), affording the corresponding product 5.
The analytical data for compounds 3 and 5 (available in the Supplementary Materials) are as follows.
2-(2-(benzo[d]thiazol-2-yl)-2,2-difluoro-1-phenylethyl)malononitrile (3a): Prepared according to the general procedure A as described above in an 81% yield (82 mg) as a yellow solid. Mp. 137.7–138.1 °C. 1H NMR (400 MHz, CDCl3) δ 8.13 (d, J = 8.0 Hz, 1H), 7.91 (d, J = 8.0 Hz, 1H), 7.61–7.57 (m, 1H), 7.53–7.48 (m, 3H), 7.38–7.36 (m, 3H), 5.02 (d, J = 8.0 Hz, 1H), 4.62–4.54 (m, 1H) ppm. 13C NMR (101 MHz, CDCl3) δ 161.45 (t, J = 33.3 Hz, 1C), 152.11, 134.94, 130.08, 129.65, 129.33, 127.16, 127.13, 124.45, 122.07, 117.60 (t, J = 249.4 Hz, 1C), 111.04, 110.77, 51.54 (t, J = 23.2 Hz, 1C), 24.67 (t, J = 4.0 Hz, 1C) ppm. 19F NMR (376 MHz, CDCl3) δ −87.11 (d, J = 270.7 Hz, 1F), −94.67 (d, J = 270.7 Hz, 1F) ppm. HRMS (ESI): exact mass calculated for C18H11F2N3S [M+H]+, 340.0705; found, 340.0715.
2-(2-(benzo[d]thiazol-2-yl)-2,2-difluoro-1-(p-tolyl)ethyl)malononitrile (3b): Prepared according to the general procedure A as described above in an 86% yield (91 mg) as a yellow solid. Mp. 120.4–120.7 °C. 1H NMR (400 MHz, CDCl3) δ 8.13 (d, J = 8 Hz, 1H), 7.91 (d, J = 8 Hz, 1H), 7.60–7.56 (m, 1H), 7.52–7.48 (m, 1H), 7.36 (d, J = 8 Hz, 2H), 7.16 (d, J = 8 Hz, 2H), 4.98 (d, J = 4 Hz, 1H), 4.50–4.58 (m, 1H), 2.30 (s, 3H) ppm. 13C NMR (101 MHz, CDCl3) δ 161.59 (t, J = 33.3 Hz, 1C), 152.14, 140.15, 134.95, 130.03, 129.47, 127.12, 127.06, 126.19, 124.44, 122.06, 117.67 (t, J = 249.4 Hz, 1C), 111.14, 110.86, 51.26 (t, J = 23.7 Hz, 1C), 24.75 (t, J = 4.0 Hz, 1C), 21.15 ppm. 19F NMR (376 MHz, CDCl3) δ −86.87 (d, J = 266.9 Hz, 1F), −94.78, −95.13 (d, J = 266.9 Hz, 1F) ppm. HRMS (ESI): exact mass calculated for C19H13F2N3S [M+H]+, 354.0862; found, 354.0871.
2-(1-([1,1′-biphenyl]-4-yl)-2-(benzo[d]thiazol-2-yl)-2,2-difluoroethyl)malononitrile (3c): Prepared according to the general procedure A as described above in a 90% yield (112 mg) as a yellow solid. Mp. 140.1–140.4 °C. 1H NMR (400 MHz, CDCl3) δ 8.14 (d, J = 8 Hz, 1H), 7.91 (d, J = 8 Hz, 1H), 7.60–7.58 (m, 5H), 7.55–7.48 (m, 3H), 7.43–7.40 (m, 2H), 7.36–7.32 (m, 1H), 5.05 (d, J = 4 Hz, 1H), 4.67–4.59 (m, 1H) ppm. 13C NMR (101 MHz, CDCl3) δ 161.44 (t, J = 33.3 Hz, 1C), 152.12, 142.77, 139.60, 134.97, 130.08, 128.81, 127.91, 127.86, 127.18, 127.15, 127.04, 124.46, 122.10, 117.61 (t, J = 249.4 Hz, 1C), 111.08, 110.81, 51.24 (t, J = 23.2 Hz, 1C), 24.72 (t, J = 4.0 Hz, 1C) ppm. 19F NMR (376 MHz, CDCl3) δ −86.85 (d, J = 270.7 Hz, 1F), −94.61 (d, J = 270.7 Hz, 1F) ppm. HRMS (ESI): Exact mass calculated for C24H15 F2N3S [M+H]+, 416.1017; found, 416.1028.
2-(2-(benzo[d]thiazol-2-yl)-1-(4-bromophenyl)-2,2-difluoroethyl)malononitrile (3d): Prepared according to the general procedure A as described above in a 66% yield (82 mg) as a yellow solid. Mp. 133.1–133.4 °C. 1H NMR (400 MHz, CDCl3) δ 8.12 (d, J = 8 Hz, 1H), 7.94 (d, J = 12 Hz, 1H), 7.62–7.58 (m, 1H), 7.55–7.51 (m, 3H), 7.40 (d, J = 12 Hz, 2H), 5.01 (d, J = 4.0 Hz, 1H), 4.62–4.54 (m, 1H) ppm. 13C NMR (101 MHz, CDCl3) δ 161.09 (t, J = 33.3 Hz, 1C), 152.04, 134.93, 132.62, 131.29, 128.55, 128.22, 127.28, 124.75, 124.47, 122.14, 117.34 (t, J = 249.4 Hz, 1C), 110.82, 110.54, 50.90 (t, J = 24.2 Hz, 1C), 24.52 (t, J = 4.0 Hz, 1C) ppm. 19F NMR (376 MHz, CDCl3) δ −86.79 (d, J = 270.7 Hz, 1F), −95.01 (d, J = 270.7 Hz, 1F) ppm. HRMS (ESI): exact mass calculated for C18H1079BrF2N3S [M+H]+, 417.9809; found, 417.9820.
2-(2-(benzo[d]thiazol-2-yl)-1-(4-chlorophenyl)-2,2-difluoroethyl)malononitrile (3e): Prepared according to the general procedure A as described above in an 83% yield (92 mg) as a yellow solid. Mp. 168.8–169.1 °C. 1H NMR (400 MHz, CDCl3) δ 8.12 (d, J = 8 Hz, 1H), 7.935 (d, J = 4 Hz, 1H), 7.58–7.62 (m, 1H), 7.53–7.55 (m, 1H), 7.36 (d, J = 8 Hz, 2H), 5.01 (d, J = 8 Hz, 1H), 4.55–4.63 (m, 1H) ppm. 13C NMR (101 MHz, CDCl3) δ 161.08 (t, J = 33.3 Hz, 1C), 152.04, 136.44, 134.93, 131.05, 129.66, 127.73, 127.70, 127.28, 124.46, 122.14, 117.41 (t, J = 249.4 Hz, 1C), 110.84, 110.56, 50.83 (t, J = 23.7 Hz, 1C), 24.59 (t, J = 4.0 Hz, 1C) ppm. 19F NMR (376 MHz, CDCl3) δ −86.84 (d, J = 270.7 Hz, 1F), −95.08 (d, J = 270.7 Hz, 1F) ppm. HRMS (ESI): exact mass calculated for C18H1035ClF2N3S [M+H]+, 374.0316; found, 374.0325.
2-(2-(benzo[d]thiazol-2-yl)-1-(3,5-dimethylphenyl)-2,2-difluoroethyl)malononitrile (3f): Prepared according to the general procedure A as described above in a 75% yield (82 mg) as a yellow solid. Mp. 116.4–116.8 °C. 1H NMR (400 MHz, CDCl3) δ 8.13 (d, J = 8 Hz, 1H), 7.91 (d, J = 8 Hz, 1H), 7.60–7.56 (m, 1H), 7.52–7.48 (m, 1H), 7.07 (s, 2H), 6.99 (s, 1H), 4.96 (d, J =8 Hz, 1H), 4.52–4.43 (m, 1H), 2.27 (s, 6H) ppm. 13C NMR (101 MHz, CDCl3) δ 161.72 (t, J = 33.3 Hz, 1C), 152.17, 138.92, 135.02, 131.79, 129.14, 127.33, 127.12, 127.07, 124.45, 122.05, 117.64 (t, J = 249.4 Hz, 1C), 111.13, 110.86, 51.46 (t, J = 23.7 Hz, 1C), 24.76 (t, J = 4.0 Hz, 1C), 21.26 ppm. 19F NMR (376 MHz, CDCl3) δ −87.55 (d, J = 270.7 Hz, 1F), −93.60 (d, J = 270.7 Hz, 1F) ppm. HRMS (ESI): exact mass calculated for C20H15 F2N3S [M+H]+, 368.1021; found, 368.1028.
4-(2-(benzo[d]thiazol-2-yl)-1-(4-bromophenyl)-2,2-difluoroethyl)-2,6-di-tert-butylphenol (5a): Prepared according to the general procedure B as described above in a 65% yield (72 mg) as a white solid. Mp. 154–156 °C. 1H NMR (400 MHz, CDCl3) δ 8.12 (d, J = 8.0 Hz, 1H), 7.86 (d, J = 4.0 Hz, 1H), 7.54–7.51 (m, 1H), 7.46–7.41 (m, 5H), 7.15 (s, 2H), 5.19–5.10 (m, 2H), 1.33 (s, 18H) ppm. 13C NMR (101 MHz, CDCl3) δ 165.04 (t, J = 34.3 Hz, 1C), 153.18, 152.58, 136.14, 135.70, 135.03, 131.56, 131.29, 126.62, 126.35, 126.25, 126.19, 124.05, 121.80, 121.40, 119.80 (t, J = 249.5 Hz, 1C), 56.98 (t, J = 23.2 Hz, 1C), 34.24, 30.12 ppm. 19F NMR (376 MHz, CDCl3) δ −92.37 (d, J = 259.4 Hz, 1F), −93.23 (d, J = 259.4 Hz, 1F) ppm. HRMS (ESI): exact mass calculated for C29H2979BrF2NOS [M-H]−, 556.1127; found, 556.1137.
4-(2-(benzo[d]thiazol-2-yl)-1-(4-chlorophenyl)-2,2-difluoroethyl)-2,6-di-tert-butylphenol (5b): Prepared according to the general procedure B as described above in a 66% yield (68 mg) as a yellow solid. Mp. 134–136 °C. 1H NMR (400 MHz, CDCl3) δ 8.11 (d, J = 8.0 Hz, 1H), 7.85 (d, J = 8.0 Hz, 1H), 7.54–7.50 (m, 1H), 7.46–7.41 (m, 3H), 7.24–7.22 (m, 3H), 7.14 (s, 2H), 5.19–5.10 (m, 2H), 1.32 (s, 18H) ppm. 13C NMR (101 MHz, CDCl3) δ 165.07 (t, J = 33.3 Hz, 1C), 153.17, 152.58, 135.69, 135.61, 135.03, 133.18, 130.94, 128.61, 126.62, 126.36, 126.25, 124.05, 121.80, 119.86 (t, J = 249.5 Hz, 1C), 56.92 (t, J = 23.3 Hz, 1C), 34.25, 30.12 ppm. 19F NMR (376 MHz, CDCl3) δ −92.41 (d, J = 255.7 Hz, 1F), −93.34 (d, J = 255.7 Hz, 1F) ppm. HRMS (ESI): exact mass calculated for C29H2935ClF2NOS [M-H]−, 512.1632; found, 512.1650.
4-(2-(benzo[d]thiazol-2-yl)-2,2-difluoro-1-phenylethyl)-2,6-di-tert-butylphenol (5c): Prepared according to the general procedure B as described above in a 62% yield (59 mg) as a yellow solid. Mp. 140–142 °C. 1H NMR (400 MHz, CDCl3) δ 8.13 (d, J = 8.0 Hz, 1H), 7.84 (d, J = 8.0 Hz, 1H), 7.55–7.50 (m, 3H), 7.44–7.40 (m, 1H), 7.31–7.28 (m, 2H), 7.24–7.21 (m, 3H), 5.21–5.11 (m, 2H), 1.33 (s, 18H) ppm. 13C NMR (101 MHz, CDCl3) δ 165.45 (t, J = 33.3 Hz, 1C), 153.04, 152.61, 137.03, 135.53, 135.07, 129.62, 128.43, 127.23, 126.75, 126.51, 126.49, 126.11, 124.04, 121.74, 120.07 (t, J = 250.5 Hz, 1C), 57.66 (t, J = 23.2 Hz, 1C), 34.23, 30.14 ppm. 19F NMR (376 MHz, CDCl3) δ −92.34 (d, J = 259.4 Hz, 1F), −93.20 (d, J = 259.4 Hz, 1F) ppm. HRMS (ESI): exact mass calculated for C29H30F2NOS [M-H]−, 478.2022; found, 478.2046.
4-(2-(benzo[d]thiazol-2-yl)-2,2-difluoro-1-(p-tolyl)ethyl)-2,6-di-tert-butylphenol (5d): Prepared according to the general procedure B as described above in a 66% yield (65 mg) as a yellow solid. Mp. 149–150 °C. 1H NMR (400 MHz, CDCl3) δ 8.13 (d, J = 8.0 Hz, 1H), 7.84 (d, J = 8.0 Hz, 1H), 7.54–7.50 (m, 1H), 7.44–7.40 (m, 3H), 7.21 (s, 2H), 7.10 (d, J = 8.0 Hz, 2H), 5.17–5.08 (m, 2H), 2.29 (s, 3H), 1.34 (s, 18H) ppm. 13C NMR (101 MHz, CDCl3) δ 165.57 (t, J = 33.3 Hz, 1C), 152.98, 152.62, 136.83, 135.48, 135.09, 133.98, 129.46, 129.16, 126.97, 126.48, 126.42, 126.06, 124.03, 121.74, 120.11 (t, J = 249.5 Hz, 1C), 57.33 (t, J = 23.2 Hz, 1C), 34.23, 30.16, 21.00 ppm. 19F NMR (376 MHz, CDCl3) δ −92.74, −94.34 ppm. HRMS (ESI): exact mass calculated for C30H32F2NOS [M-H]−, 492.2178; found, 492.2184.
4-(2-(benzo[d]thiazol-2-yl)-2,2-difluoro-1-(4-methoxyphenyl)ethyl)-2,6-di-tert-butylphenol (5e): Prepared according to the general procedure B as described above in a 65% yield (66 mg) as a yellow solid. Mp. 127–128 °C. 1H NMR (400 MHz, CDCl3) δ 8.13 (d, J = 8.0 Hz, 1H), 7.84 (d, J = 8.0 Hz, 1H), 7.54–7.50 (m, 1H), 7.46–7.40 (m, 3H), 7.22 (s, 2H), 6.83 (d, J = 8.0 Hz, 2H), 5.17–5.08 (m, 2H), 3.75 (s, 3H), 1.35 (s, 18H) ppm. 13C NMR (101 MHz, CDCl3) δ 165.56 (t, J = 34.5 Hz, 1C), 158.61, 152.96, 152.59, 135.50, 135.06, 130.69, 129.08, 127.05, 126.48, 126.37, 126.06, 123.99, 121.73, 120.12 (t, J = 249.5 Hz, 1C), 113.78, 56.87 (t, J = 23.2 Hz, 1C), 55.06, 34.22, 30.15 ppm. 19F NMR (376 MHz, CDCl3) δ −92.41 (d, J = 259.4 Hz, 1F), −93.40 (d, J = 259.4 Hz, 1F) ppm. HRMS (ESI): exact mass calculated for C30H32F2NO2S [M-H]−, 508.2127; found, 508.2154.
4-(1-([1,1′-biphenyl]-4-yl)-2-(benzo[d]thiazol-2-yl)-2,2-difluoroethyl)-2,6-di-tert-butylphenol (5f): Prepared according to the general procedure B as described above in a 63% yield (70 mg) as a yellow solid. Mp. 149–150 °C. 1H NMR (400 MHz, CDCl3) δ 8.14 (d, J = 8.0 Hz, 1H), 7.85 (d, J = 8.0 Hz, 1H), 7.62 (d, J = 8.0 Hz, 2H), 7.58–7.51 (m, 5H), 7.45–7.39 (m, 3H), 7.34–7.31 (m, 3H), 7.24 (s, 2H), 5.26–5.12 (m, 2H), 1.35 (s, 18H) ppm. 13C NMR (101 MHz, CDCl3) δ 165.40 (t, J = 33.3 Hz, 1C), 153.10, 152.63, 140.54, 139.93, 136.09, 135.60, 135.10, 129.99, 128.69, 127.24, 127.12, 126.96, 126.70, 126.54, 126.49, 126.14, 124.05, 121.77, 120.09 (t, J = 249.5 Hz, 1C), 57.37 (t, J = 23.2 Hz, 1C), 34.26, 30.16 ppm. 19F NMR (376 MHz, CDCl3) δ −92.18 (d, J = 259.4 Hz, 1F), −93.16 (d, J = 259.4 Hz, 1F) ppm. HRMS (ESI): exact mass calculated for C35H34F2NOS [M-H]−, 554.2335; found, 554.2356.
4-(1-(4-bromophenyl)-2-(6-chlorobenzo[d]thiazol-2-yl)-2,2-difluoroethyl)-2,6-di-tert-butylphenol (5g): Prepared according to the general procedure B as described above in a 62% yield (73 mg) as a yellow solid. Mp. 153–155 °C. 1H NMR (400 MHz, CDCl3) δ 8.01 (d, J = 8.0 Hz, 1H), 7.83 (m, 1H), 7.49 (d, J = 8.0 Hz, 1H), 7.43–7.37 (m, 4H), 7.15 (s, 2H), 5.15–5.06 (m, 2H), 1.34 (s, 18H) ppm. 13C NMR (101 MHz, CDCl3) δ 165.51 (d, J = 34.3 Hz, 1H), 153.24, 151.08, 136.19, 135.95, 135.77, 132.41, 131.61, 131.24, 127.62, 126.31, 126.01, 124.82, 121.50, 121.39, 119.60 (t, J = 249.5 Hz, 1H), 56.95 (t, J = 23.2 Hz, 1H), 34.26, 30.14 ppm. 19F NMR (376 MHz, CDCl3) δ −92.36 (d, J = 259.4 Hz, 1F), −93.25 (d, J = 259.4 Hz, 1F) ppm. HRMS (ESI): exact mass calculated for C29H28BrClF2NOS [M-H]−, 590.0737; found, 590.0748.
4. Conclusions
In conclusion, copper-catalyzed radical 1,4- or 1,6-conjugate additions of 2-Bromodifluoromethylbenzo-1,3-thiazole with arylidenemalonitriles or para-quinone methides have been developed for the synthesis of benzothiazolyldifluoroalkyl containing (di)arylmethine compounds in moderate to excellent yields. The reaction performed well on a gram scale, indicating that it was a practical tool for the synthesis of gem-difluoroalkylatated (di)arylmethine derivatives. More catalytic reactions involving 2-bromodifluoromethylbenzo-1,3-thiazoles will be reported in due course.
Supplementary Materials
The following supporting information can be downloaded at: https://www.mdpi.com/article/10.3390/catal14110777/s1, Figure S1. 1H-NMR spectrum of the reaction product of 3a; Figure S2. 13C-NMR spectrum of the reaction product of 3a; Figure S3. 19F-NMR spectrum of the reaction product of 3a; Figure S4. 1H-NMR spectrum of the reaction product of 3b; Figure S5. 13C-NMR spectrum of the reaction product of 3b; Figure S6. 19F-NMR spectrum of the reaction product of 3b; Figure S7. 1H-NMR spectrum of the reaction product of 3c; Figure S8. 13C-NMR spectrum of the reaction product of 3c; Figure S9. 19F-NMR spectrum of the reaction product of 3c; Figure S10. 1H-NMR spectrum of the reaction product of 3d; Figure S11. 13C-NMR spectrum of the reaction product of 3d; Figure S12. 19F-NMR spectrum of the reaction product of 3d; Figure S13. 1H-NMR spectrum of the reaction product of 3e; Figure S14. 13C-NMR spectrum of the reaction product of 3e; Figure S15. 19F-NMR spectrum of the reaction product of 3e; Figure S16. 1H-NMR spectrum of the reaction product of 3f; Figure S17. 13C-NMR spectrum of the reaction product of 3e; Figure S18. 19F-NMR spectrum of the reaction product of 3f; Figure S19. 1H-NMR spectrum of the reaction product of 5a; Figure S20. 13C-NMR spectrum of the reaction product of 5a; Figure S21. 19F-NMR spectrum of the reaction product of 5a; Figure S22. 1H-NMR spectrum of the reaction product of 5b; Figure S23. 13C-NMR spectrum of the reaction product of 5b; Figure S24. 19F-NMR spectrum of the reaction product of 5b; Figure S25. 1H-NMR spectrum of the reaction product of 5c; Figure S26. 13C-NMR spectrum of the reaction product of 5c; Figure S27. 19F-NMR spectrum of the reaction product of 5c; Figure S28. 1H-NMR spectrum of the reaction product of 5d; Figure S29. 13C-NMR spectrum of the reaction product of 5d; Figure S30. 19F-NMR spectrum of the reaction product of 5d; Figure S31. 1H-NMR spectrum of the reaction product of 5e; Figure S32. 13C-NMR spectrum of the reaction product of 5e; Figure S33. 19F-NMR spectrum of the reaction product of 5e; Figure S34. 1H-NMR spectrum of the reaction product of 5f; Figure S35. 13C-NMR spectrum of the reaction product of 5f; Figure S36. 19F-NMR spectrum of the reaction product of 5f; Figure S37. 1H-NMR spectrum of the reaction product of 5g; Figure S38. 13C-NMR spectrum of the reaction product of 5g; Figure S39. 19F-NMR spectrum of the reaction product of 5g.
Author Contributions
Prepared the compound and ran the spectra, J.X., Y.C. and R.X.; designed the study, analyzed the data, and wrote the paper, F.L.; helped to discuss the experimental results and process, J.L. All authors have read and agreed to the published version of the manuscript.
Funding
We are grateful for financial support from the Jiangxi Provincial Education Department Scientific Research Foundation (No. GJJ190434), National Natural Science Foundation of China (21961014), and Jiangxi Provincial Key Laboratory of Functional Crystalline Materials Chemistry (2024SSY05161).
Data Availability Statement
Data are contained within the article and Supplementary Materials.
Conflicts of Interest
The authors declare no conflicts of interest.
References
- Ma, J.-A.; Cahard, D. Asymmetric fluorination, trifluoromethylation, and perfluoroalkylation reactions. Chem. Rev. 2004, 104, 6119–6146. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- O’Hagan, D. Understanding organofluorine chemistry. An introduction to the C-F bond. Chem. Soc. Rev. 2008, 37, 308–319. [Google Scholar] [CrossRef] [Scilit]
- Zhang, C.-P.; Chen, Q.-Y.; Guo, Y.; Xiao, J.-C.; Gu, Y.-C. Progress in fluoroalkylation of organic compounds via sulfinatodehalogenation initiation system. Chem. Soc. Rev. 2012, 41, 4536–4559. [Google Scholar] [CrossRef] [Scilit]
- Wang, J.; Sánchez-Roselló, M.; Aceña, J.L.; del Pozo, C.; Sorochinsky, A.E.; Fustero, S.; Soloshonok, V.A.; Liu, H. Fluorine in pharmaceutical industry: Fluorine-containing drugs introduced to the market in the last decade (2001–2011). Chem. Rev. 2014, 114, 2432–2506. [Google Scholar] [CrossRef] [Scilit]
- Yang, X.; Wu, T.; Phipp, R.J.; Toste, F.D. Advances in catalytic enantioselective fluorination, mono-, di-, and trifluoromethylation, and trifluoromethylthiolation reactions. Chem. Rev. 2015, 115, 826–870. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Dong, D.-Q.; Yang, H.; Shi, J.-L.; Si, W.-J.; Wang, Z.-L.; Xu, X.-M. Promising reagents for difluoroalkylation. Org. Chem. Front. 2020, 7, 2538–2575. [Google Scholar] [CrossRef] [Scilit]
- Ojima, I. Fluorine in Medicinal Chemistry and Chemical Biology; Wiley: Berlin, Germany, 2009. [Google Scholar]
- Bey, P.; Gerhart, F.; Dorsselaer, V.V.; Danzin, C. a-(Fluorornet hy1)dehydroornit hine and a-(Fluorornet hy1)dehydroputre an alogues as irreversible inhibitors of ornithine decarboxylase. J. Med. Chem. 1983, 26, 1551–1556. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Pourquier, P.; Gioffre, C.; Kohlhagen, G.; Urasaki, Y.; Goldwasser, F.; Hertel, L.W.; Yu, S.; Pon, R.T.; Gmeiner, W.H.; Pommier, Y. Gemcitabine (2′,2′-difluoro-2′-deoxycytidine), an antimetabolite that poisons topoisomerase I. Clin. Cancer Res. 2002, 8, 2499–2504. [Google Scholar]
- Ando, K.; Koike, F.; Kondo, F.; Takayama, H. An improved synthesis of 24,24-difluoro-1α,25-dihydroxyvitamin D3 from readily available vitamin D2. Chem. Pharm. Bull. 1995, 43, 189–192. [Google Scholar] [CrossRef] [Scilit]
- Uoto, K.; Ohsuki, S.; Takenoshita, H.; Ishiyama, T.; Iimura, S.; Hirota, Y.; Mitsui, I.; Terasawa, H.; Soga, T. Synthesis and structure-activity relationships of novel 2′,2′-difluoro analogues of docetaxel. Chem. Pharm. Bull. 1997, 45, 1793–1804. [Google Scholar] [CrossRef] [Scilit]
- Nie, J.; Guo, H.-C.; Cahard, D.; Ma, J.-A. Asymmetric construction of stereogenic carbon centers featuring a trifluoromethyl group from prochiral trifluoromethylated substrates. Chem. Rev. 2011, 111, 455–529. [Google Scholar] [CrossRef] [Scilit]
- Hu, J.; Zhang, W.; Wang, F. Selective difluoromethylation and monofluoromethylation reactions. Chem. Commun. 2009, 7465–7478. [Google Scholar] [CrossRef] [Scilit]
- Liu, Y.-L.; Yu, J.-S.; Zhou, J. Catalytic asymmetric construction of stereogenic carbon centers that feature a gem-difluoroalkyl group. Asian J. Org. Chem. 2013, 2, 194–206. [Google Scholar] [CrossRef] [Scilit]
- Lin, J.-H.; Xiao, J.-C. Recent advances in asymmetric fluorination and fluoroalkylation reactions via organocatalysis. Tetrahedron Lett. 2014, 55, 6147–6155. [Google Scholar] [CrossRef] [Scilit]
- Ni, C.; Zhu, L.; Hu, J. Advances in transition-metal-mediated di- and monofluoroalkylations. Acta Chim. Sin. 2015, 73, 90–115. [Google Scholar] [CrossRef] [Scilit]
- Qing, F.-L.; Liu, X.-Y.; Ma, J.-A.; Shen, Q.; Song, Q.; Tang, P. A fruitful decade of organofluorine chemistry: New reagents and reactions. CCS Chem. 2022, 4, 2518–2549. [Google Scholar] [CrossRef] [Scilit]
- Burkholder, C.R.; Dolbier, W.R., Jr.; Médebielle, M. The syntheses of nonnucleoside, HIV-1 reverse transcriptase inhibitors containing a CF2 group The SRN1 reactions of 2-(bromodifluoromethyl)benzoxazole with the anions derived from heterocyclic thiols and phenolic compounds. J. Fluor. Chem. 2000, 102, 369–376. [Google Scholar] [CrossRef] [Scilit]
- Jiang, H.; Yuan, S.; Cai, Y.; Wan, W.; Zhu, S.; Hao, J. A facile preparation of 2-bromodifluoromethyl benzo-1,3-diazoles and its application in the synthesis of gem-difluoromethylene linked aryl ether compounds. J. Fluor. Chem. 2012, 133, 167–170. [Google Scholar] [CrossRef] [Scilit]
- Burkholder, C.R.; Dolbier Jr, W.R.; Médebielle, M. Synthesis and reactivity of halogeno-difluoromethyl aromatics heterocycles: Application to synthesis of gem-difluorinated bioactive compounds. J. Fluor. Chem. 2001, 109, 39–48. [Google Scholar] [CrossRef] [Scilit]
- Jiang, H.; Lu, W.; Yang, K.; Ma, G.; Xu, M.; Li, J.; Yao, J.; Wan, W.; Deng, H.; Wu, S.; et al. Enhancement of neighbouring group participation in Cu0 promoted cross-coupling gem-difluoromethylenation of aryl/alkenyl halides with 1,3-azolic difluoromethyl bromides. Chem.-Eur. J. 2014, 20, 10084–10092. [Google Scholar] [CrossRef] [Scilit]
- Ma, G.; Wan, W.; Hu, Q.; Jiang, H.; Wang, J.; Zhu, S.; Hao, J. Highly effective copper-mediated gem-difluoromethylenation of arylboronic acids. Chem. Commun. 2014, 50, 7527–7530. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Jiang, H.; Xu, M.; Lu, W.; Tian, W.; Wan, W.; Chen, Y.; Deng, H.; Wu, S.; Hao, J. Direct gem-difluoromethylenation of sp3-hybridized carbon center through copper-mediated radical/radical cross-coupling for the construction of a CH2-CF2 linkage. Chem. Commun. 2015, 51, 15756–15759. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Xiao, Y.-L.; Zhang, B.; Feng, Z.; Zhang, X. Heteroaryldifluoromethylation of organoborons catalyzed by palladium: Facile access to aryl(heteroaryl)difluoromethanes. Org. Lett. 2014, 16, 4822–4825. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Gu, J.-W.; Zhang, X. Palladium-catalyzed difluoroalkylation of isocyanides: Access to difluoroalkylated phenanthridine derivatives. Org. Lett. 2015, 17, 5384–5387. [Google Scholar] [CrossRef] [Scilit]
- Xu, C.; Yang, Z.-F.; An, L.; Zhang, X. Nickel-catalyzed difluoroalkylation-alkylation of enamides. ACS Catal. 2019, 9, 8224–8229. [Google Scholar] [CrossRef] [Scilit]
- Xu, H.; Wang, D.; Chen, Y.; Wan, W.; Deng, H.; Ma, K.; Wu, S.; Hao, J.; Jiang, H. Copper-catalyzed gem-difluoromethylenation of C(sp2)-H bonds of alkenes. Org. Chem. Front. 2017, 4, 1239–1243. [Google Scholar] [CrossRef] [Scilit]
- Zhang, T.; Chen, B.; Wang, W.; Zhang, Q.; Wang, P.; Wan, W.; Deng, H.; Hao, J.; Jiang, H. Copper-promoted aryldifluoromethylenation of N-arylacrylamides to 3-Benzodiazolyldifluoromethylene substituted 2-oxindoles. Asian J. Org. Chem. 2019, 8, 671–674. [Google Scholar] [CrossRef] [Scilit]
- Zhang, Q.; Wang, W.; Wang, P.; Deng, H.; Jiang, H. Copper-catalyzed aryldifluoromethylenation of N-arylacrylamides to synthesis of the diheterocyclic compounds linked by gem-difluoromethylene moiety. Tetrahedron 2019, 75, 130778. [Google Scholar] [CrossRef] [Scilit]
- Da, Y.; Han, S.; Du, X.; Liu, S.; Liu, L.; Li, J. Copper(I)-catalyzed oxydifluoroalkylation of alkenes: A route to functionalization of lactones. Org. Lett. 2018, 20, 5149–5152. [Google Scholar] [CrossRef] [Scilit]
- Li, X.; Hu, K.; Zhang, J.; Jiang, H. Cu0-promoted Truce-Smiles rearrangement for aryl-difluoromethylenation of C=C bonds via a reductive radical-polar crossover process. J. Org. Chem. 2024, 89, 13947–13952. [Google Scholar] [CrossRef] [Scilit]
- Malapit, C.A.; Reeves, J.T.; Busacca, C.A.; Howell, A.R.; Senanayake, C.H. Rhodium-catalyzed transnitrilation of aryl boronic acids with dimethylmalononitrile. Angew. Chem. Int. Ed. 2016, 55, 326–330. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Malapit, C.A.; Caldwell, D.R.; Luvaga, I.K.; Reeves, J.T.; Volchkov, I.; Gonnella, N.C.; Han, Z.S.; Busacca, C.A.; Howell, A.R.; Senanayake, C.H. Rhodium-catalyzed addition of aryl boronic acids to 2,2-disubstituted malononitriles. Angew. Chem. Int. Ed. 2017, 56, 6999–7002. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Peter, M.G. Chemical modifications of biopolymers by quinones and quinone methides. Angew. Chem. Int. Ed. Engl. 1989, 28, 555–570. [Google Scholar] [CrossRef] [Scilit]
- Abrams, P.; Freeman, R.; Anderström, C.; Mattiasson, A. Tolterodine, a new antimuscarinic agent: As effective but better tolerated than oxybutynin in patients with an overactive bladder. Br. J. Urol. 1998, 81, 801–810. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- An, R.-B.; Jeong, G.-S.; Kim, Y.-C. Flavonoids from the heartwood of dalbergia odorifera and their protective effect on glutamate-induced oxidative injury in HT22 cells. Chem. Pharm. Bull. 2008, 56, 1722–1724. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Davidson, S.J.; Barker, D. Synthesis of various lignans via the rearrangements of 1,4-diarylbutane-1,4-diols. Tetrahedron Lett. 2015, 56, 4549–4553. [Google Scholar] [CrossRef] [Scilit]
- Yu, J.-S.; Liao, F.-M.; Gao, W.-M.; Liao, K.; Zuo, R.-L.; Zhou, J. Michael addition catalyzed by chiral secondary amine phosphoramide using fluorinated silyl enol ethers: Formation of quaternary carbon stereocenters. Angew. Chem. Int. Ed. 2015, 54, 7381–7385. [Google Scholar] [CrossRef] [Scilit]
- Hao, Y.-J.; Hu, X.-S.; Yu, J.-S.; Zhou, F.; Zhou, Y.; Zhou, J. An efficient Fe(III)-catalyzed 1,6-conjugate addition of para-quinone methides with fluorinated silyl enol ethers toward b,b-diaryl afluorinated ketones. Tetrahedron 2018, 74, 7395–7398. [Google Scholar] [CrossRef] [Scilit]
- Ke, M.; Song, Q. Copper-catalyzed 1,6-hydrodifluoroacetylation of para-quinone methides at ambient temperature with bis(pinacolato)diboron as reductant. Adv. Synth. Catal. 2017, 359, 384–389. [Google Scholar] [CrossRef] [Scilit]
- Zhao, Y.-N.; Luo, Y.-C.; Wang, Z.-Y.; Xu, P.-F. A new approach to access difluoroalkylated diarylmethanes via visible-light photocatalytic cross-coupling reactions. Chem. Commun. 2018, 54, 3993–3996. [Google Scholar] [CrossRef] [Scilit]
- Qu, C.-H.; Song, G.-T.; Tang, D.-Y.; Shao, J.-W.; Li, H.-Y.; Xu, Z.-G.; Chen, Z.-Z. Microwave-assisted copper catalysis of α-difluorinated gem-diol toward difluoroalkyl radical for hydrodifluoroalkylation of para-quinone methides. J. Org. Chem. 2020, 85, 12785–12796. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Xu, R.; Liao, F.; Cai, Y.; Liu, J. A convenient synthesis of 2-bromodifluoromethyl benzo-1,3-thiazoles. Chin. J. Syn. Chem. 2020, 30, 588–592. [Google Scholar]
- Xu, R.; Cai, Y.; Liao, F.; Liu, J. 2-[Difluoro(phenylselenyl)methyl]benzo-1,3-thiazole. Molbank 2022, 2022, 1450. [Google Scholar] [CrossRef] [Scilit]
- Xu, R.; Liao, F.; Cai, Y.; Liu, J. Copper-catalyzed aryl selenylation of 2-bromodifluoromethylbenzo-1,3-thi(x)azoles with diaryl diselenides. Tetrahedron 2022, 128, 133118. [Google Scholar] [CrossRef] [Scilit]
- Petrier, C.; Dupuy, C.; Luche, J.L. Conjugate additions to α,β-unsaturated carbonyl compounds in aqueous media. Tetrahedron Lett. 1986, 27, 3149–3152. [Google Scholar] [CrossRef] [Scilit]
- Suárez, R.M.; Sestelo, J.P.; Sarandeses, L.A. Diastereoselective conjugate addition to chiral α,β-unsaturated carbonyl systems in aqueous media: An enantioselective entry to α- and γ-hydroxy acids and α-amino acids. Chem.-Eur. J. 2003, 9, 4179–4187. [Google Scholar] [CrossRef] [Scilit]
- Shin, J.-A.; Kim, J.; Lee, H.; Ha, S.; Lee, H.-Y. Cu(OTf)2-promoted 1,4-addition of alkyl bromides to dehydroalanine. J. Org. Chem. 2019, 84, 4558–4565. [Google Scholar] [CrossRef] [Scilit]
- Gugkaeva, Z.T.; Smol’yakov, A.F.; Maleev, V.I.; Larionov, V.A. A general asymmetric synthesis of artificial aliphatic and perfluoroalkylated α-amino acids by Luche’s cross-electrophile coupling reaction. Org. Biomol. Chem. 2021, 19, 5327–5332. [Google Scholar] [CrossRef] [Scilit]

Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2024 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license (https://creativecommons.org/licenses/by/4.0/).