


Abstract
A new bi-triazole precursor, N-(1-azido-2-(azidomethyl)butan-2-yl)-4-methylbenzenesulfonamide, was synthesized in two steps from 2-amino-2-ethyl-1,3-propanediol, with an overall yield of 80%. The chemical structures of the products obtained were established based on 1D and 2D NMR, IR spectroscopy, and elemental analysis.
1. Introduction
The synthesis of new molecular units with drug-like properties remains a major concern in the field of drug chemistry, particularly in the fight against cancer and infectious diseases. The preparation of bi-triazole systems constitutes a matter of urgency for researchers [1], mainly due to their broad spectrum of applications. Thus, in recent years, triazoles have increasingly been targeted due to their interesting antiproliferative [2], antifungal [3], antidiabetic [4], antibacterial [5], anticancer [6], antioxidants, and anti-inflammatory properties [7]. They are also used as corrosion inhibitors [8].
Given these observations and the continuity of our previous work concerning the synthesis of derivatives of heterocyclic amino acids and their precursors [9,10,11,12,13,14,15], we were interested in performing the synthesis and characterization of a new precursor of the bi-triazole systems.
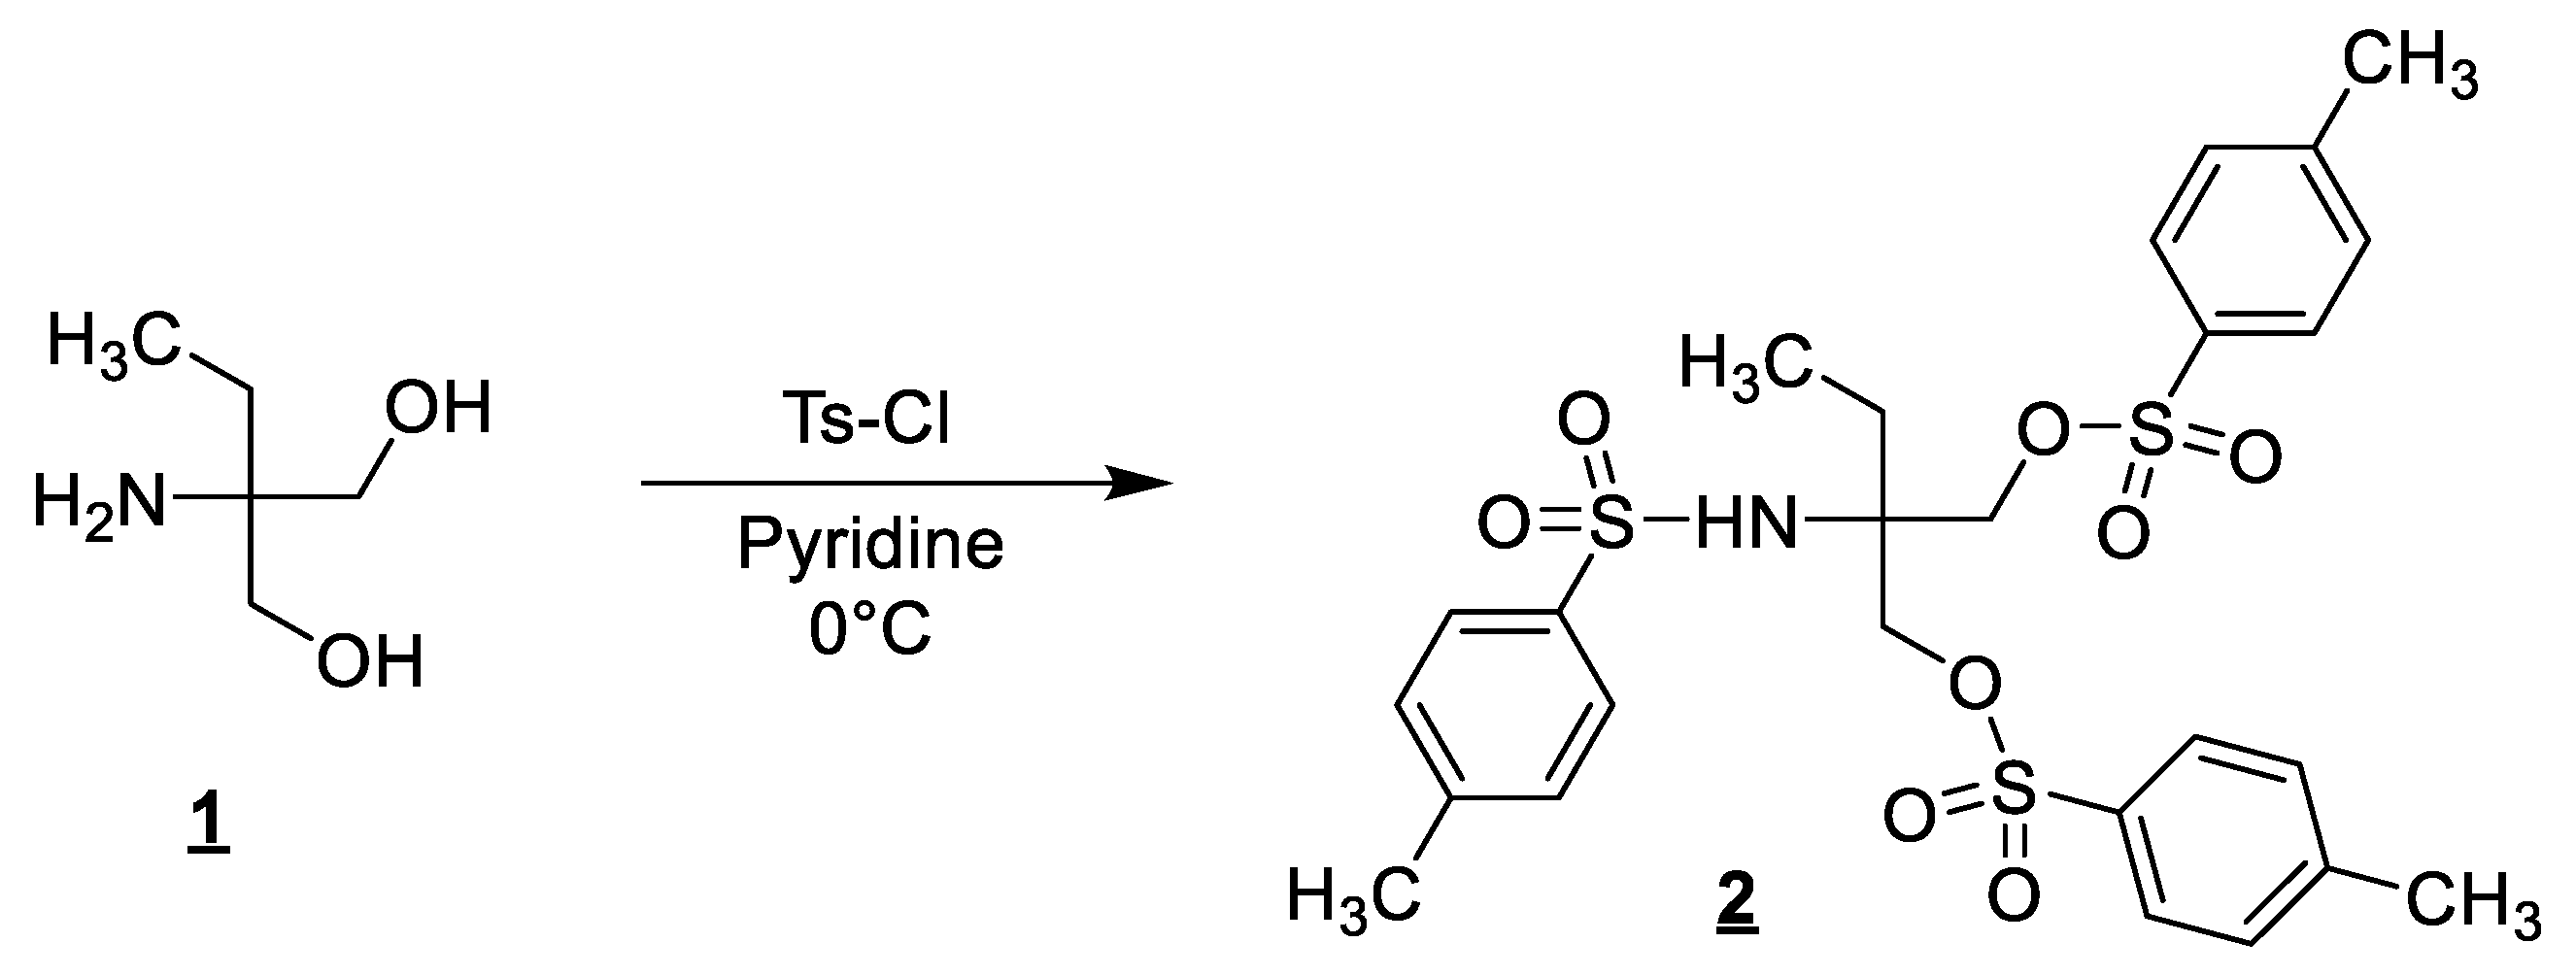
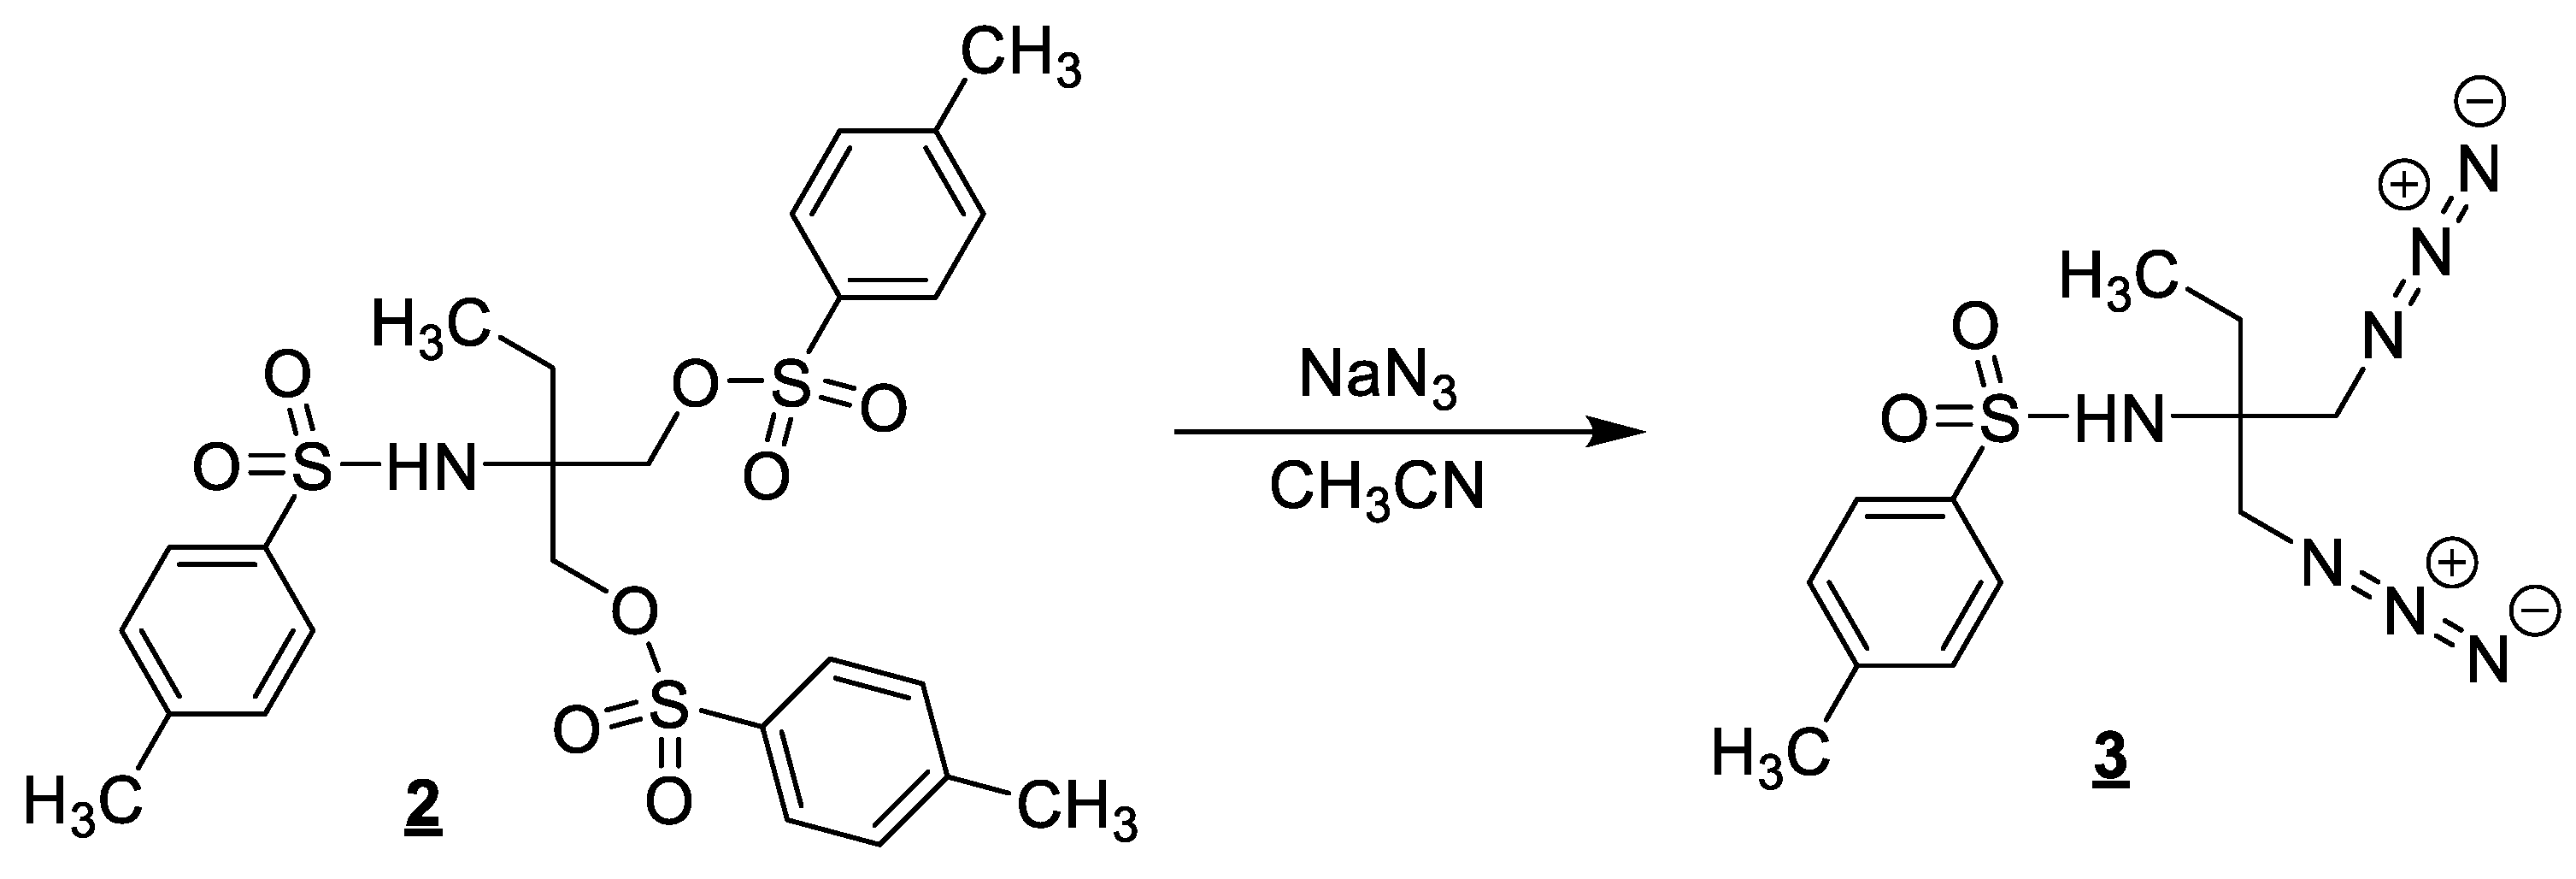
We report, in this article, on the synthesis of a new precursor of bi-triazole compounds obtained in two steps: The first step included the tosylation of 2-amino-2-ethyl-1,3-propanediol (1) to obtain the di-O-tosyl compound, namely 2-ethyl-2-((4-methylphenyl)sulfonamido)propane-1,3-diylbis(4-methylbenzenesulfonate) (2). this was followed by the substitution of the O-tosyl groups of the latter with -N3, which allowed us to obtain the desired compound (3) in a good yield. The compounds obtained were characterized by 1D and 2D NMR, IR, and elemental analysis (Supplementary Materials).
2. Results
The action of tosyl chloride in pyridine on 2-amino-2-ethyl-1,3-propanediol (1) leads to the compound 2-ethyl-2-((4-methylphenyl)sulfonamido)propane-1,3-diyl bis(4-methylbenzenesulfonate) (2), with a yield of pure product of 60% (Scheme 1). Then, the obtained product is subjected to the action of sodium azide in acetonitrile (Scheme 2).
Scheme 1.
Synthesis of compound (2).
Scheme 2.
Synthesis of compound (3).
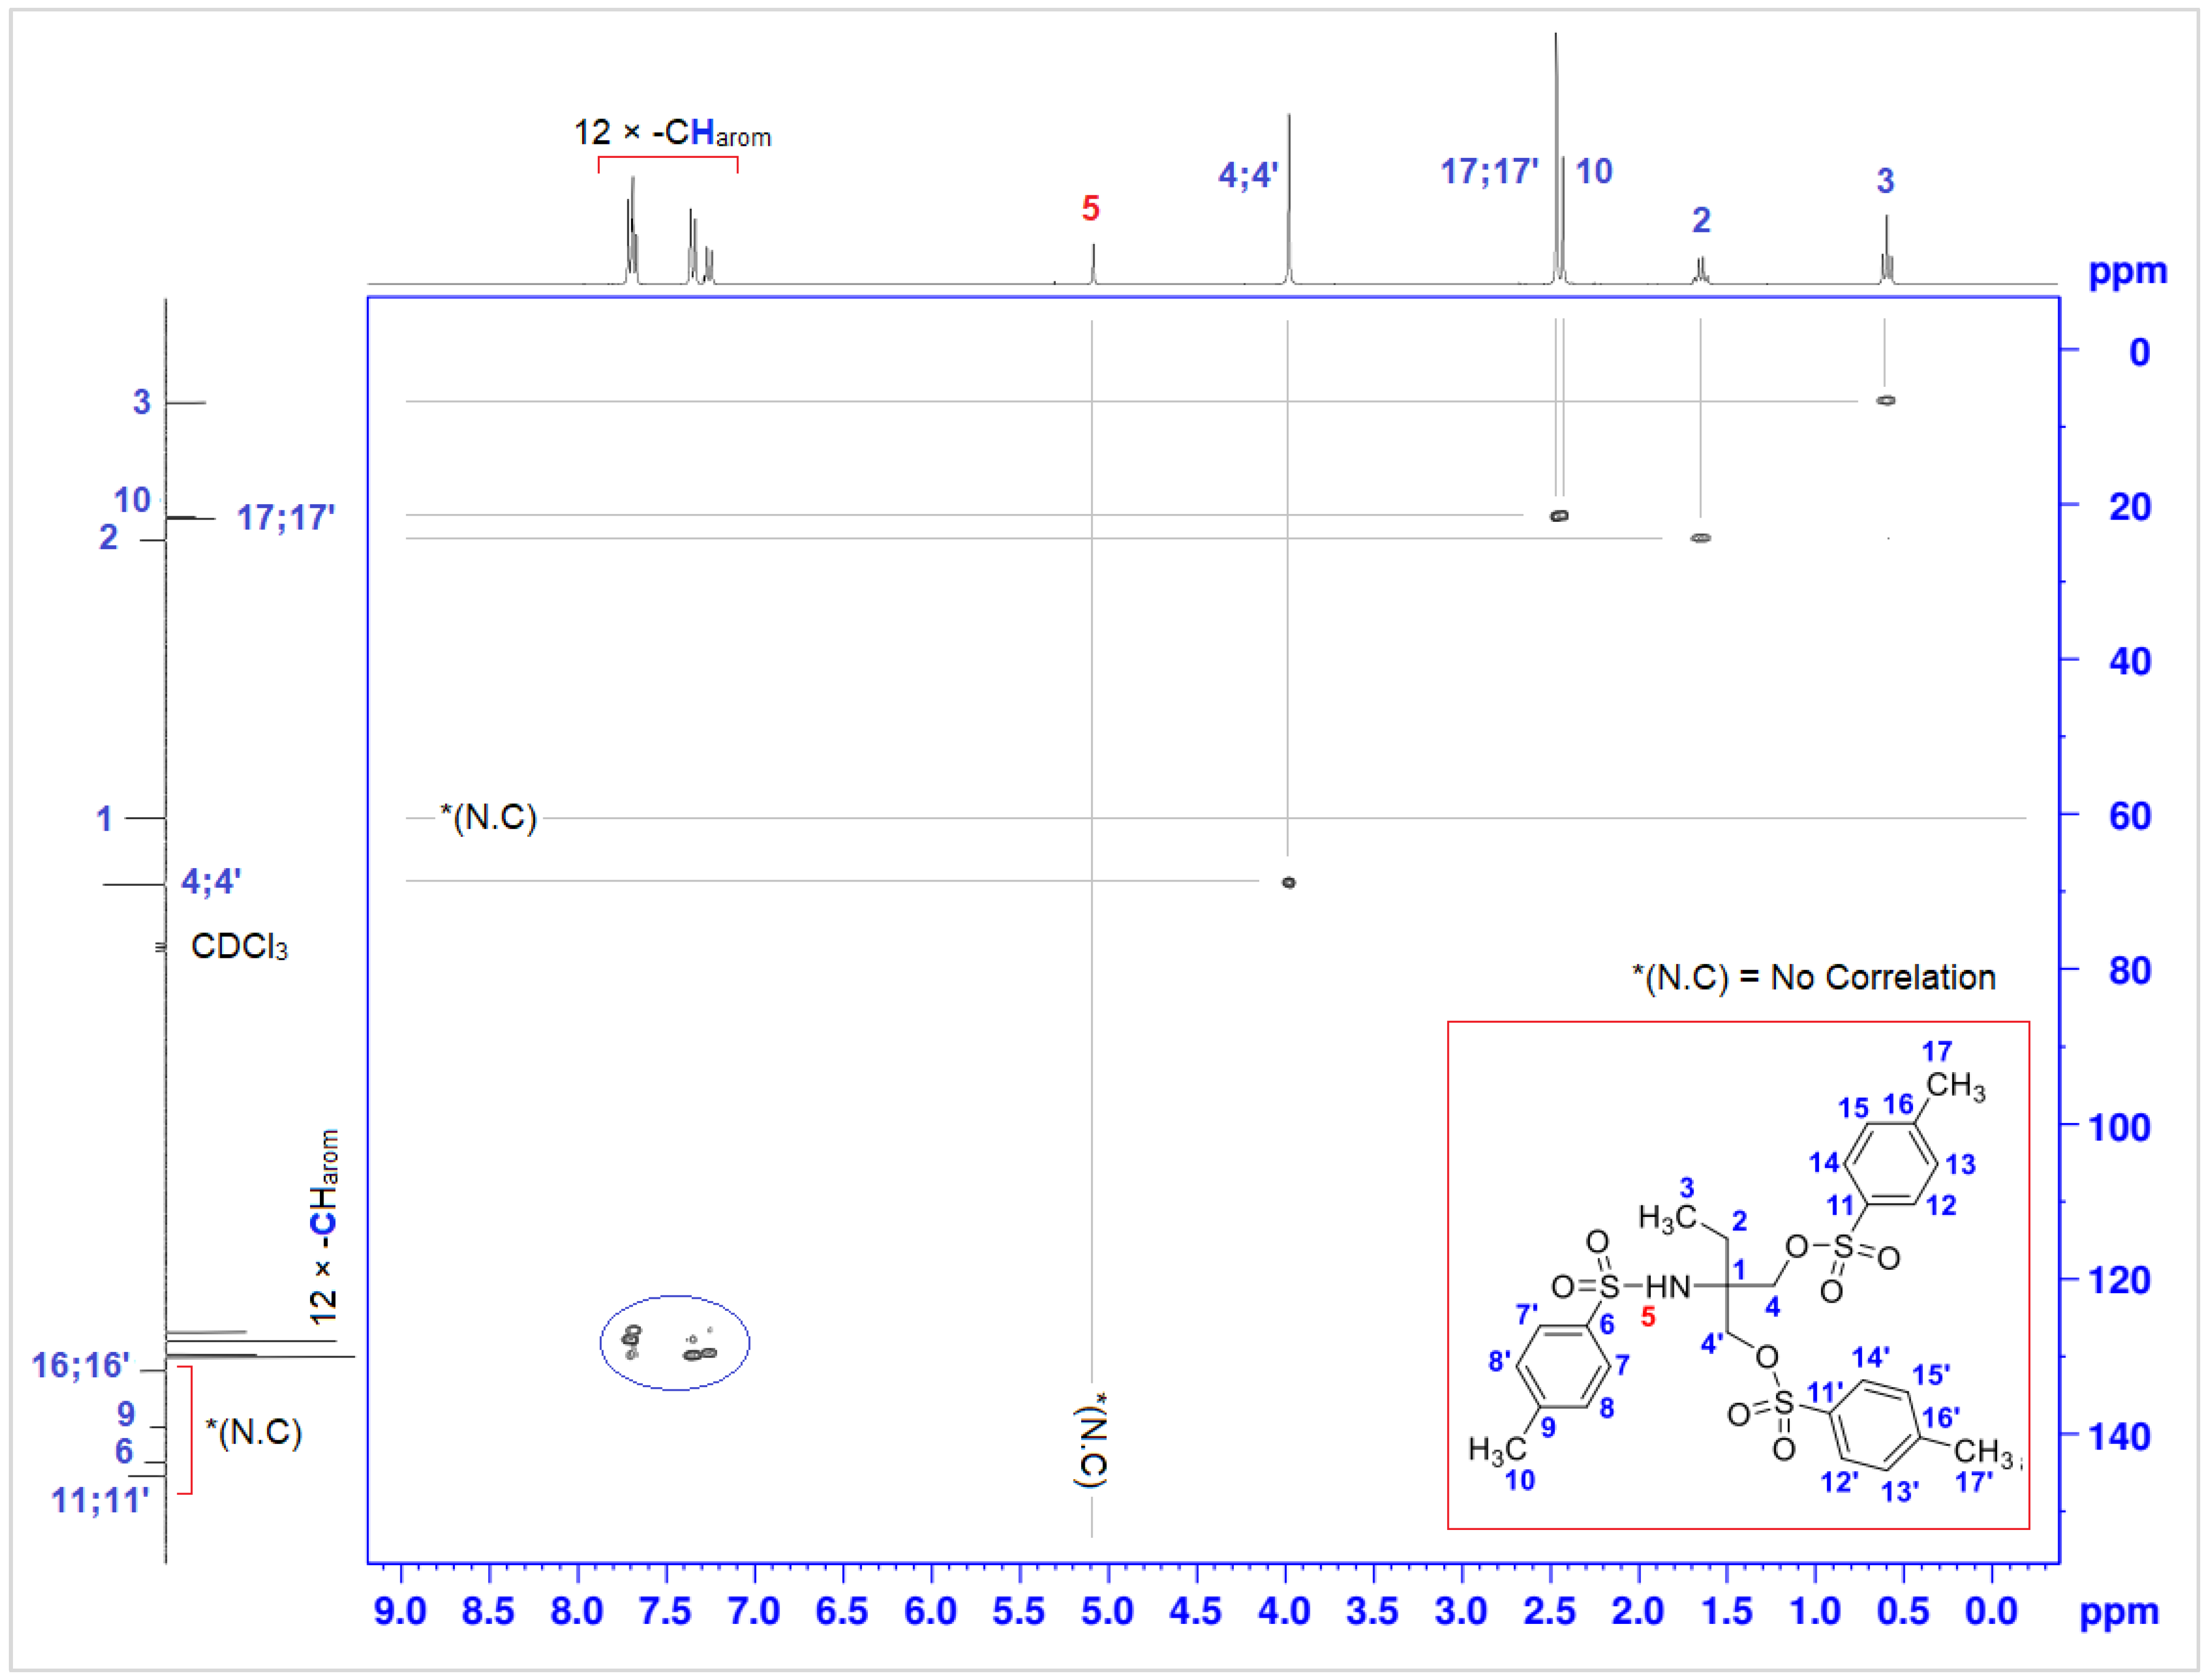
The structures of the compounds obtained were elucidated using standard spectroscopic analytical methods (1D and 2D NMR) and infrared spectroscopy. The 1H NMR spectrum of compound (2) shows the presence of a singlet at 2.43 ppm, corresponding to the three methyl protons bound to -NHTs. In addition, the methyl group protons bonded to -OTs resonate at 2.46 ppm as a singlet. In addition, the 12 aromatic protons appear as four doublets at around 7.24–7.71 ppm (J = 8.1 Hz). Moreover, for the 13C NMR spectrum, which shows the presence of a signal at 21.6 ppm, corresponding to the methyl carbon linked to -NHTs, there is a signal at 21.7 ppm relating to the two carbons of the methyl groups linked to -OTs.
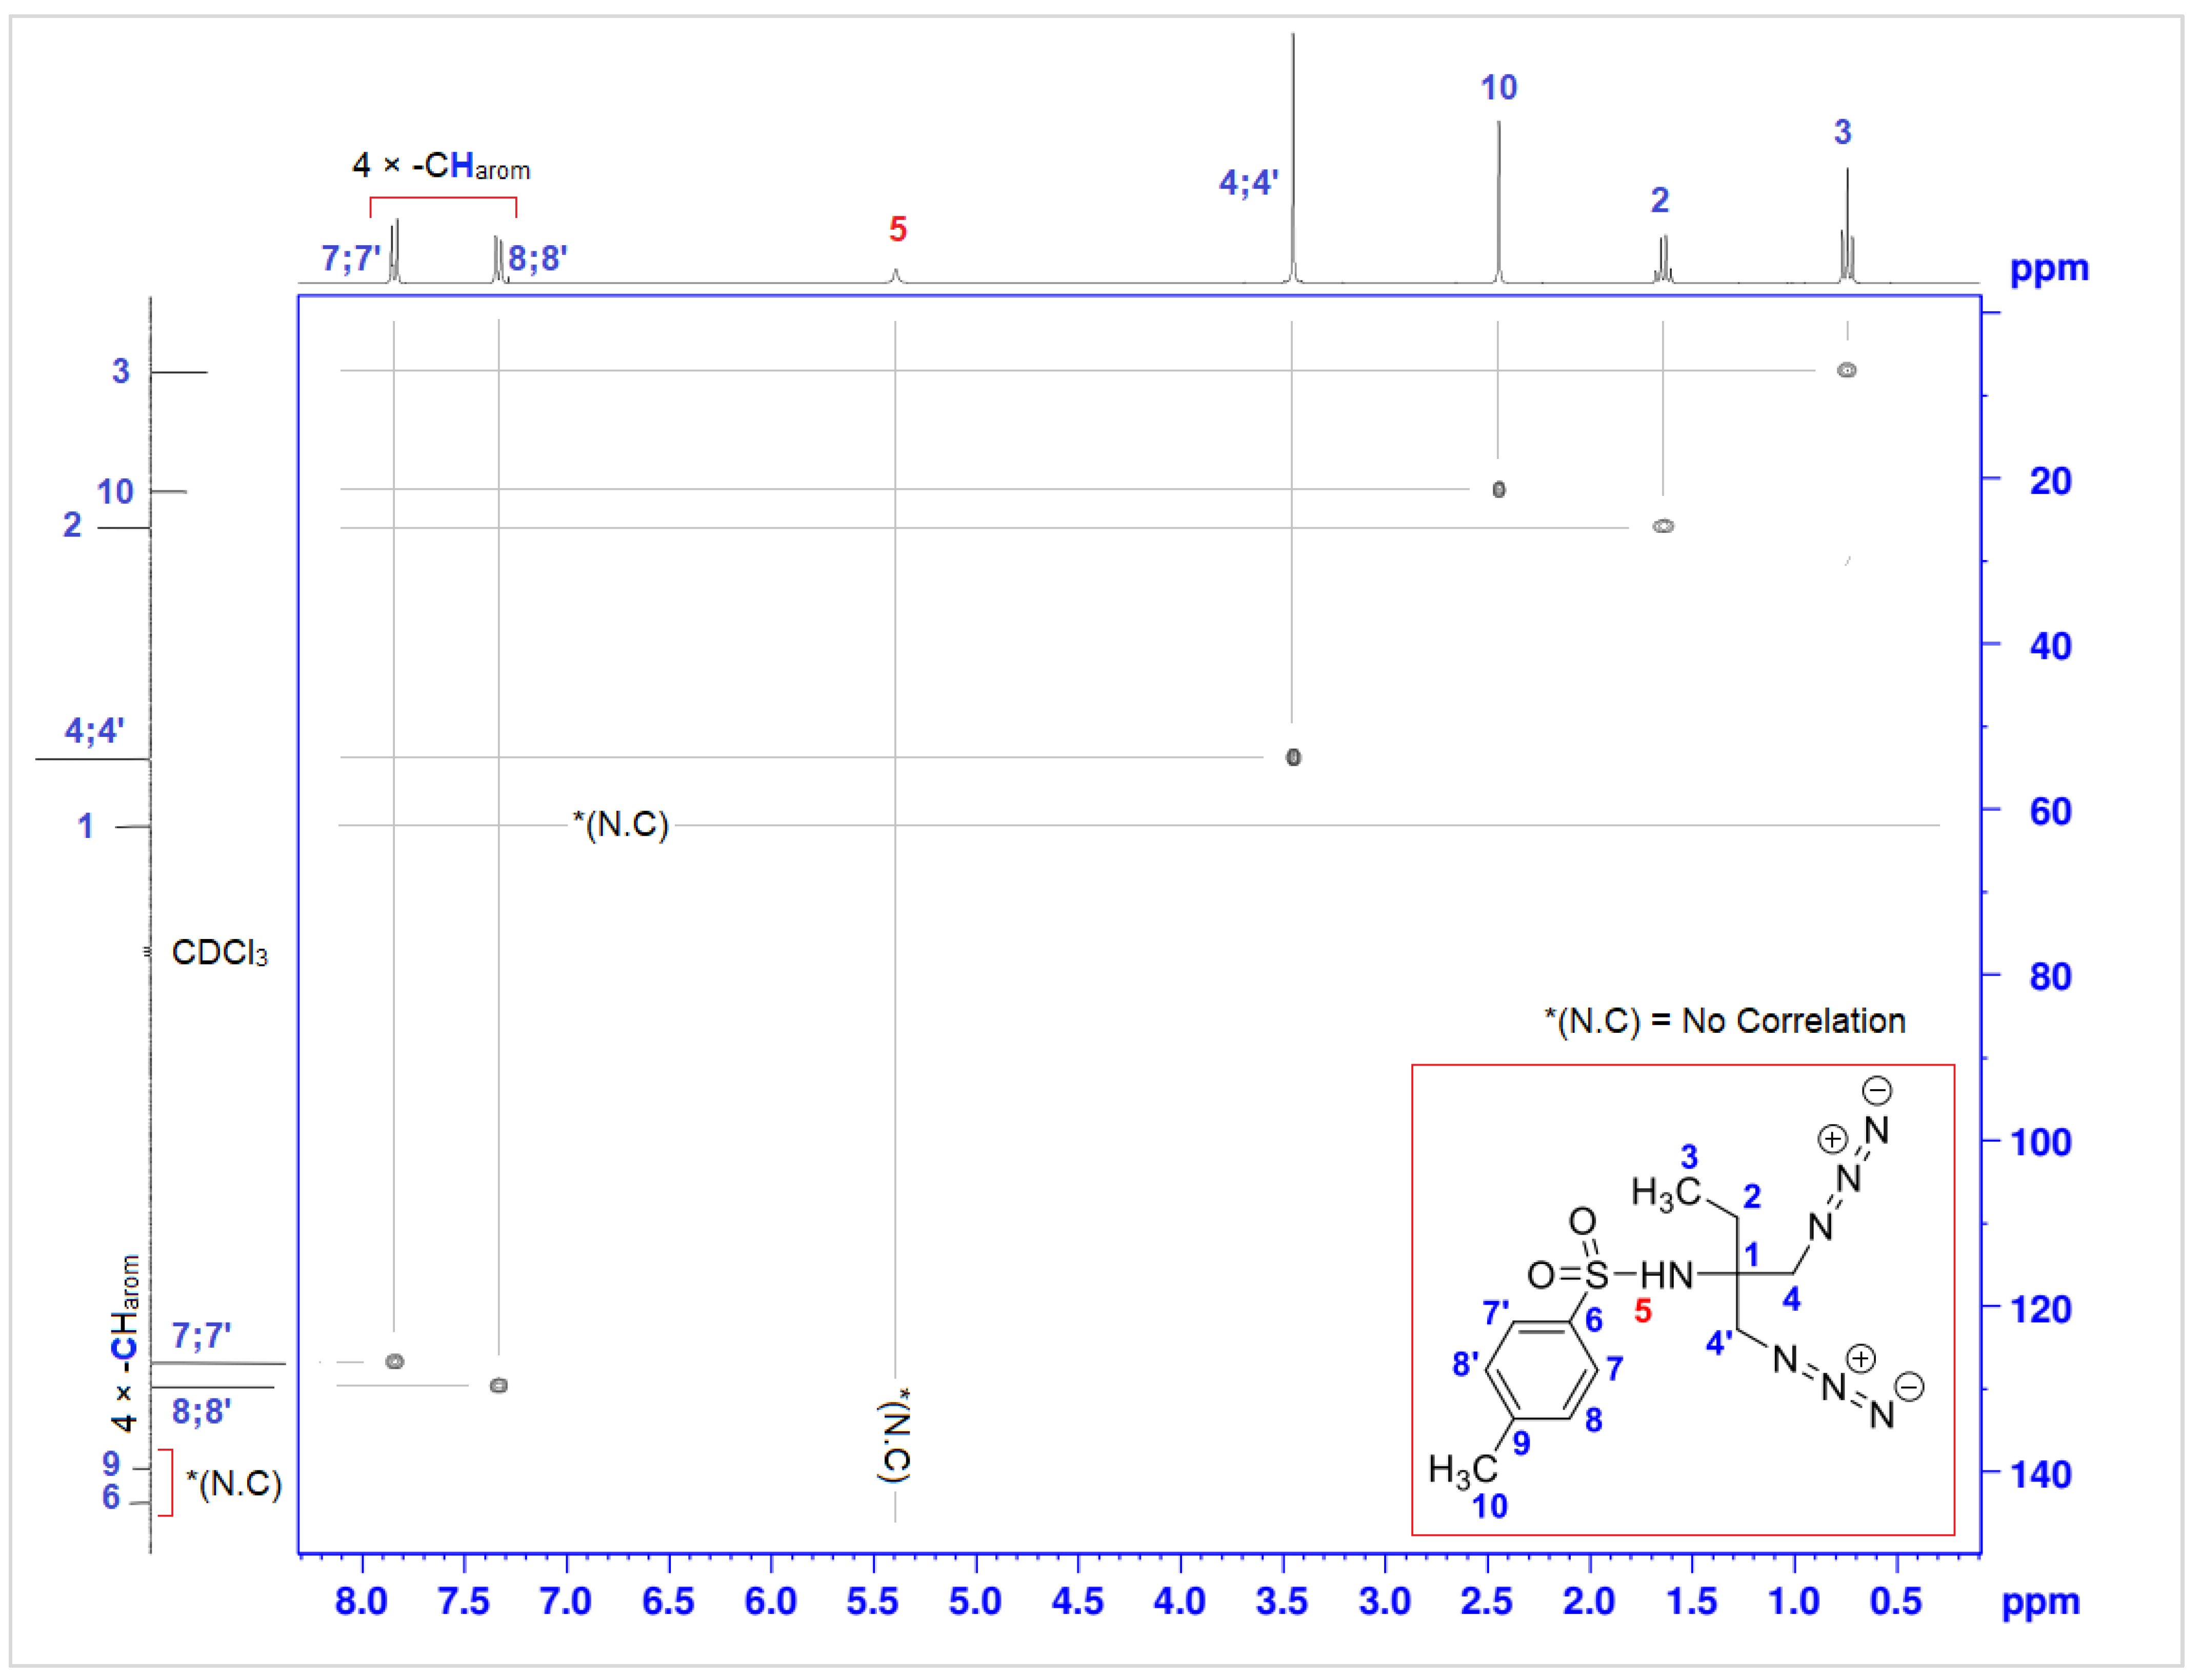
Furthermore, the 1H-NMR spectrum of compound (3) shows a singlet at 2.44 ppm, attributed to the three protons of the methyl group bonded to -NHTs. It also shows two doublets at 7.31–7.34 and 7.82–7.85 ppm, corresponding only to the four aromatic protons (J = 8.1 Hz). On the other hand, the 13C NMR spectrum of this compound exhibits a signal at 21.5 ppm, attributed to the methyl group carbon bonded to -NHTs. The precise assignment of the chemical shifts of the protons and carbons is presented in Table 1 and Table 2. The interpretation of the 2D HSQC NMR spectra (Figure 1 and Figure 2) of the two compounds (2) and (3) showed a perfect correlation between proton–proton and proton–carbon 13.
Table 1.
1H (300 MHz) and 13C (75 MHz) NMR spectral data for compound (2) in CDCl3, including results obtained by homonuclear 2D-shift-correlated and heteronuclear 2D-shift-correlated HMBC. Chemical shifts (δ in ppm) and coupling constants (J in Hz).
Table 2.
1H (300 MHz) and 13C (75 MHz) NMR spectral data for compound (3) in CDCl3, including results obtained by homonuclear 2D-shift-correlated and heteronuclear 2D-shift-correlated HMBC. Chemical shifts (δ in ppm) and coupling constants (J in Hz).
Figure 1.
2D HSQC spectrum of compound (2). “*”: No correlation.
Figure 2.
2D HSQC spectrum of compound (3). “*”: No correlation.
The IR spectrum of compound (3) shows, inter alia, a high-intensity band at 2100 cm−1, characteristic of the stretching vibrations of the azide group (-N3). Thus, in the spectrum of compound (2), we note the absence of this last band, which shows that the substitution reaction of the leaving groups -OTs by the azide groups (-N3) has taken place.
3. Materials and Methods
All solvents were purified following the standard techniques, and commercial reagents were purchased from Sigma-Aldrich (St. Louis, MO, USA). The melting point was determined using an electrothermal melting point apparatus and was uncorrected. The NMR spectra (1H and 13C) were recorded on a Bruker AM 300 spectrometer (operating at 300 MHz for 1H and at 75 MHz for 13C) (Bruker Analytische Messtechnik & GmbH, Rheinstetten, Germany). NMR data are listed in ppm and are reported relative to tetramethylsilane (1H, 13C). NMR spectroscopic data were recorded in CDCl3 using, as internal standards, the residual non-deuterated signal (δ = 7.26 ppm) for 1H NMR and the deuterated solvent signal (δ = 77.16 ppm) for the 13C NMR spectroscopy. Chemical shifts (δ) are given in ppm and coupling constants (J) are given in Hz. The following abbreviations are used for the multiplicities: s = singlet, d = doublet, t = triplet, and q = quartet. All reactions were followed by TLC. TLC analyses were carried out on 0.25 mm-thick precoated silica gel plates (Merck Fertigplatten Kieselgel 60F254), and spots were visualized under UV light or by exposure to vaporized iodine. The FT-IR spectrum was recorded using a KBr pellet on a Bruker Vertex 70 FTIR spectrometer. The elemental analysis was performed using a Flash 2000 EA 1112 Thermo Fisher Scientific-Elemental Analyzer (CNRST, Rabat, Morocco).
3.1. Synthesis of 2-Ethyl-2-((4-methylphenyl)sulfonamido)propane-1,3-diyl bis(4-methylbenzenesulfonate) (2)
To 42 mmol (5 g) of 2-amino-2-ethyl-1,3-propanediol, 10 mL of pyridine is added under stirring at 0 °C, followed by 0.21 mol (39.9 g, 5 eq) of 4-methylbenzenesulfonyl chloride (Ts-Cl), which is added in small portions. After stirring overnight, acidified water (pH between 3–4) is added to the pasty solution formed. The reaction crude is extracted with methylene chloride (3 × 30 mL). The organic phase is washed with acidified water (6 × 30 mL) and then dried and concentrated.
Yield = 75% (white solid); m.p = 132 °C; Rf = 0.36 (ethyl acetate/hexane: 1/4). 1H NMR (CDCl3, δH ppm, 300 MHz): 0.59 (3H, -CH2-CH3, t, J = 7.5 Hz); 1.65 (2H, -CH2-CH3, q, J = 7.5 Hz); 2.42 (3H, -NHTs-CH3, s); 2.46 (6H, 2 × (-OTs-CH3), s); 3.97 (4H, 2 × (-CH2-OTs), s); 5.08 (1H, -NHTs-CH3, s); 7.24–7.27; 7.66–7.69 (4 × -CHarom, 2d, J = 8.1 Hz); 7.33–7.36; 7.68–7.71 (8 × -CHarom, 2d, J = 8.1 Hz). 13C NMR (CDCl3, δC ppm, 75 MHz): 6.8 (1C, -CH2-CH3); 21.6 (1C, -NHTs-CH3); 21.7 (2C, 2 × (-OTs-CH3); 24.5 (1C, -CH2-CH3); 60.4 (1C, Cq(sp3)); 69.0 (2C, 2 × (-CH2-OTs)); 126.84–130.07 (12C, -CHarom); 131.85 (1C, -Cq(arom)-CH3); 139.12 (2C, -Cq(arom)-CH3); 143.7 (1C, -Cq(arom)-Ts-NH-); 145.4 (2C, -Cq(arom)-TsO-). IR (ν (cm−1)): 3240 (N-H(stretching)); 2960 (=CH(stretching)); 1600 (N-H(bending)); 1350–1450 (S=O(stretching)); 800–850 (=CH(bending)).
3.2. Synthesis of N-(1-azido-2-(azidomethyl)butan-2-yl)-4-methylbenzenesulfonamide (3)
To 90 mL of acetonitrile, 28 mmol (10g) of the product (2) is added with 98 mmol (6.37 g, 3.5 eq) of sodium azide. Then, the mixture is brought to reflux by stirring for 24 h at 80 °C. After the reaction, the mixture is filtered and concentrated in a vacuum. The residue obtained is washed with water (3 × 30 mL). The organic phase is dried and then concentrated. The crude reaction is purified by chromatography on a silica gel column (eluent: 10 % methylene chloride/5% ethyl acetate/85% hexane).
Yield = 80% (white solid); m.p = 159 °C; Rf = 0,08 (ethyl acetate /hexane: 1/4). 1H NMR (CDCl3, δH ppm, 300 MHz): 0.74 (3H, -CH2-CH3, t, J = 7.5 Hz); 1.64 (2H, -CH2-CH3, q, J = 7.5 Hz); 2.44 (3H, -NHTs-CH3, s); 3.45 (4H, 2 × (-CH2-N3), s); 5.39 (1H, -NHTs-CH3, s); 7.31–7.34 (2 × -CHarom, d, J = 8.1 Hz); 7.82–7.85 (2 × -CHarom, d, J = 8.1 Hz). 13C NMR (CDCl3, δC ppm, 75 Hz): 7.1 (1C, -CH2-CH3); 21.5 (1C, -NHTs-CH3); 25.9 (1C, -CH2-CH3); 53.9 (2C, 2 × (-CH2-N3)); 62.0 (1C, Cq(sp3)); 126.8 and 129.8 (4C, -CHarom); 139.6 and 143.7 (2C, -Cq(arom)). IR (ν (cm−1)): 3250 (N-H(stretching)); 2950 (=CH(stretching)); 2100 (-N3); 1600 (N-H(bending)); 1300–1350 (S=O(stretching)); 820 (=CH(bending)). Anal. Calcd. for C12H17N7O2S (%): C, 44.57; H, 5.30; N, 30.32; found (%): C, 44.68; H, 5.42; N, 30.17.
4. Conclusions
The synthesis of the title compound, N-(1-azido-2-(azidomethyl)butan-2-yl)-4-methylbenzenesulfonamide, was carried out with a good yield via the substitution reaction of the O-tosyl groups of 2-ethyl-2-((4-methylphenyl)sulfonamido)propane-1,3-diylbis(4-methylbenzenesulfonate) with the azide group (-N3). The characterization of the structures of the two products obtained (2) and (3) was carried out by 1D and 2D NMR spectroscopy, as well as IR.
Supplementary Materials
The following materials are available online, Figure S1: 1H-NMR spectrum of compound (2); Figure S2: 13C-NMR spectrum of compound (2); Figure S3: 2D-HSQC spectrum of compound (2); Figure S4: IR spectrum of compound (2); Figure S5: 1H-NMR spectrum of compound (3); Figure S6: 13C-NMR spectrum of compound (3); Figure S7: 2D-HSQC spectrum of compound (3); Figure S8: IR spectrum of compound (3).
Author Contributions
S.H. performed the experiments; H.F. and A.A. conceived and designed the experiments; Y.A. and A.A. analyzed the data and wrote the paper. All authors have read and agreed to the published version of the manuscript.
Funding
This research received no external funding.
Data Availability Statement
The data presented in this study are available in the Supplementary Materials.
Acknowledgments
This work was supported by Sidi Mohammed Ben Abdellah University (USMBA) and National Center for Scientific and Technical Research (CNRST).
Conflicts of Interest
The authors declare no conflict of interest.
References
- Dawood, K.M.; Abdel-Wahab, B.F.; Raslan, M.A. Synthesis and applications of bi- and bis-triazole systems. Arkivoc 2018, part i, 179–215. [Google Scholar] [CrossRef] [Scilit]
- Dwivedi, A.R.; Kumar, V.; Yadav, R.P.; Kumar, N.; Jangid, K.; Anand, P.; Sharma, D.K.; Barnawal, S.; Kumar, V. Design, synthesis and evaluation of 4-phenyl-1,2,3-triazole substituted pyrimidine derivatives as antiproliferative and tubulin polymerization inhibitors. J. Mol. Struct. 2022, 1267, 133592. [Google Scholar] [CrossRef] [Scilit]
- Sadeghian, S.; Emami, L.; Mojaddami, A.; Khabnadideh, S.; Faghih, Z.; Zomorodian, K.; Rashidi, M.; Rezaei, Z. 1,2,4-Triazole derivatives as novel and potent antifungal agents: Design, synthesis and biological evaluation. J. Mol. Struct. 2022, 1271, 134039. [Google Scholar] [CrossRef] [Scilit]
- Mohamed, M.A.A.; Abd Allah, O.A.; Bekhit, A.A.; Kadry, A.M.; El-Saghier, A.M.M. Synthesis and antidiabetic activity of novel triazole derivatives containing amino acids. J. Heterocycl. Chem. 2020, 57, 2365–2378. [Google Scholar] [CrossRef] [Scilit]
- Singh, G.; Singh, J.; Singh, A.; Singh, J.; Kumar, M.; Gupta, K.; Chhibber, S. Synthesis, characterization and antibacterial studies of schiff based 1,2,3-triazole bridged silatranes. J. Organomet. Chem. 2018, 871, 21–27. [Google Scholar] [CrossRef] [Scilit]
- Slaihim, M.M.; Al-Suede, F.S.R.; Khairuddean, M.; Khadeer Ahamed, M.B.; Shah Abdul Majid, A.M. Synthesis, characterization of new derivatives with mono ring system of 1,2,4-triazole scaffold and their anticancer activities. J. Mol. Struct. 2019, 1196, 78–87. [Google Scholar] [CrossRef] [Scilit]
- Rani, A.; Singh, G.; Singh, A.; Maqbool, U.; Kaur, G.; Singh, J. CuAAC-ensembled 1,2,3-triazole-linked isosteres as pharmacophores in drug discovery: Review. RSC Adv. 2020, 10, 5610–5635. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Fernandes, C.M.; Alvarez, L.X.; Escarpini dos Santos, N.; Barrios, A.C.M.; Ponzio, E.A. Green synthesis of 1-benzyl-4-phenyl-1H-1,2,3-triazole, its application as corrosion inhibitor for mild steel in acidic medium and new approach of classical electrochemical analyses. Corros. Sci. 2019, 149, 185–194. [Google Scholar] [CrossRef] [Scilit]
- Aouine, Y.; Faraj, H.; Alami, A.; El Hallaoui, A.; Elachqar, A.; Kerbal, A. Simple and efficient synthesis of racemic 2-(tert-butoxycarbon-ylamino)-2-methyl-3-(1H-1,2,4- triazol-1-yl)propanoic acid, a new derivative of β-(1,2,4-triazol-1-yl)alanine. Molecules 2011, 16, 3380–3390. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Atmani, A.; El Hallaoui, A.; El Hajji, S.; Roumestant, M.L.; Viallefont, P. From oxazolines to precursors of amino acids. Synth. Commun. 1991, 21, 2383–2390. [Google Scholar] [CrossRef] [Scilit]
- Aouine, Y.; Faraj, H.; Alami, A.; El Hallaoui, A.; EL Hajji, S.; Labriti, B.; Kerbal, A. Triheterocyclic compounds, oxazolinic precursors of biheterocyclic amino acids, part II: Phenothiazine derivatives and structural study of regioisomers through 1H-15N 2D NMR HMBC. J. Mar. Chim. Heterocycl. 2014, 13, 39–47. [Google Scholar] [CrossRef]
- Aouine, Y.; El Hallaoui, A.; Alami, A. N,N-Dibenzyl-1-(1-[(4-methyl-2-phenyl-4,5-dihydrooxazol-4-yl)methyl)]-1H-1,2,3-triazol-4-yl)methanamine. Molbank 2014, 2014, M819. [Google Scholar] [CrossRef] [Scilit]
- Hajib, S.; Alami, A.; Faraj, H.; Aouine, Y. 4-[(3,5-Dimethyl-1H-pyrazol-1-yl)methyl]-4-methyl-2-phenyl-4,5-dihydrooxazole. Molbank 2019, 2019, M1074. [Google Scholar] [CrossRef] [Scilit]
- Dioukhane, K.; Alami, A.; Aouine, Y.; El Omari, M.; El Ammari, L.; Saadi, M.; Assani, A.; Ouarsal, R. Synthesis, crystal structure and IR spectrum studies of 2-(4-methyl-2- phenyl-4,5-dihydro-oxazol-4-ylmethyl)- isoindole-1,3-dione. Mediterr. J. Chem. 2019, 9, 116–124. [Google Scholar] [CrossRef] [Scilit]
- Achamlale, S.; Alami, A.; Aouine, Y. Structure assignment of N-protected 2-(1H-1,2,3-triazol-1-yl)-glycine derivatives by chemical and spectroscopic methods. J. Mar. Chim. Heterocycl. 2019, 18, 61–69. [Google Scholar] [CrossRef]

Publisher’s Note: MDPI stays neutral with regard to jurisdictional claims in published maps and institutional affiliations. |
© 2022 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license (https://creativecommons.org/licenses/by/4.0/).