


Vibrational Entropy of Crystalline Solids from Covariance of Atomic Displacements

Abstract
1. Introduction
2. Methods
2.1. Probability Density Function
2.2. Relation to Force Constant Matrix
2.3. Quantum Harmonic Entropy
2.4. Ab Initio Methods
3. Applications
3.1. Test Cases: FCC Al and BCC Na
3.2. High Entropy Alloy: Vibrational Entropies of MoNbTaW
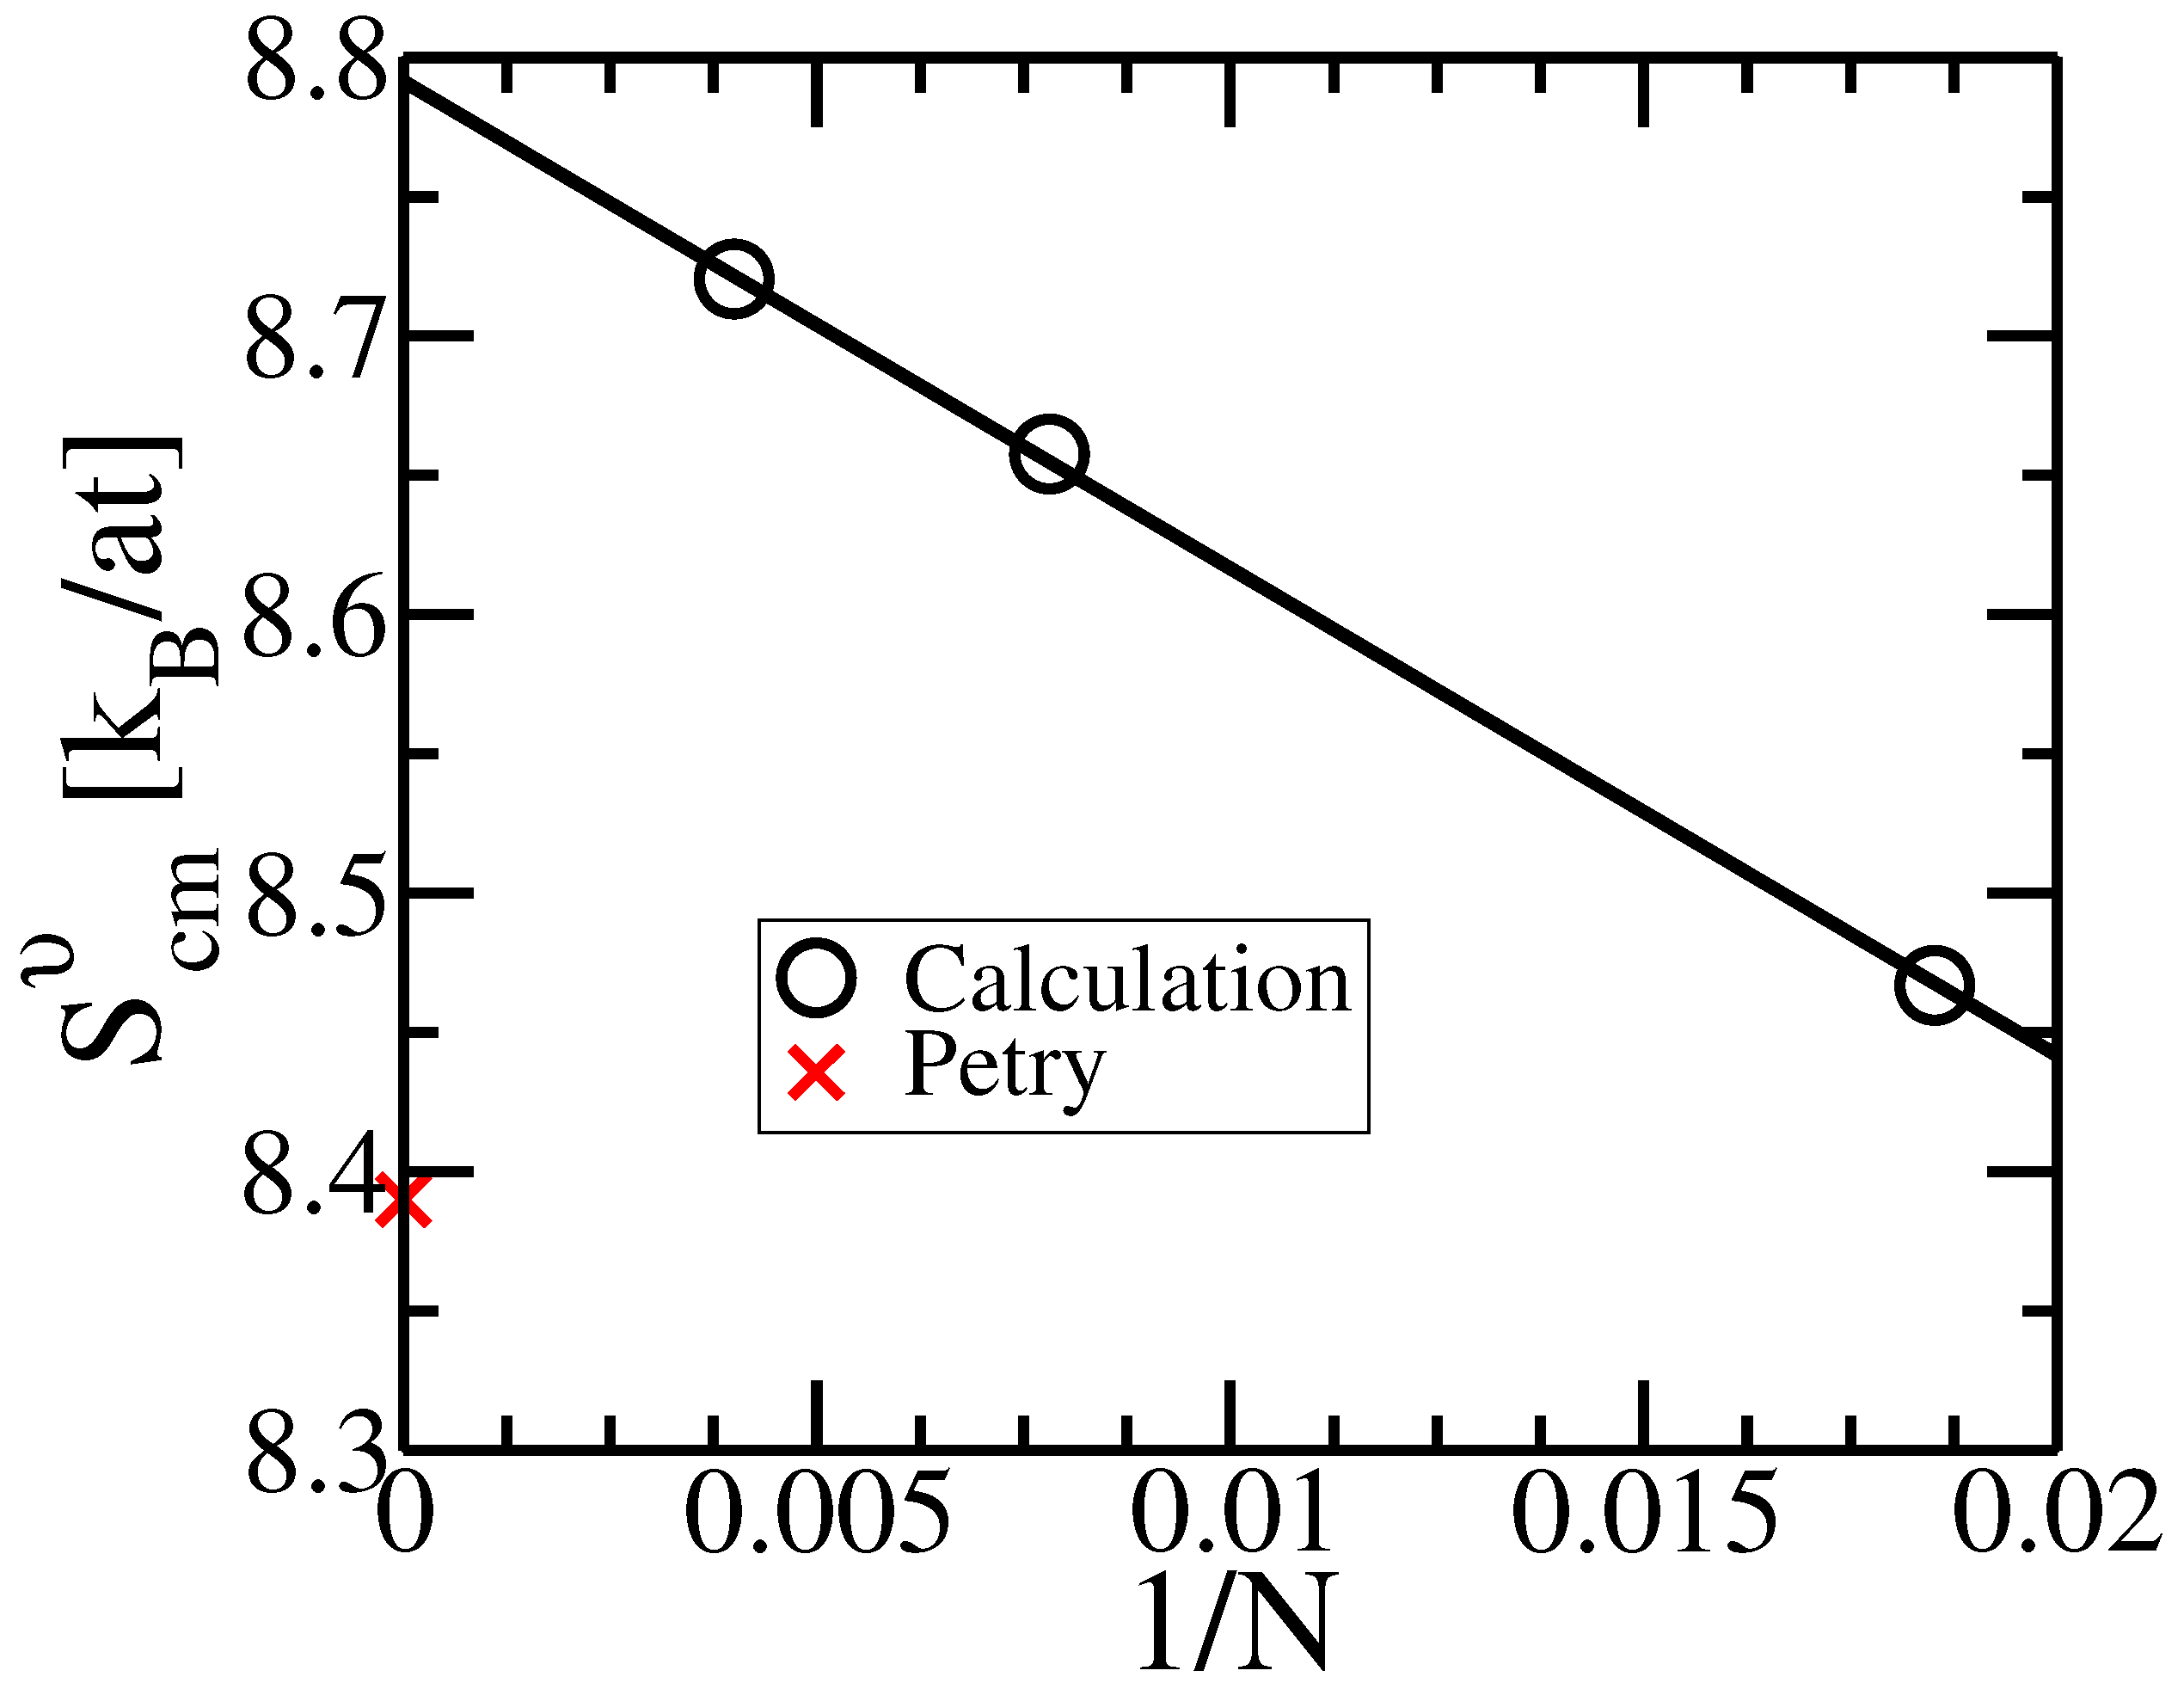
3.3. BCC to HCP Phase Transition in Titanium
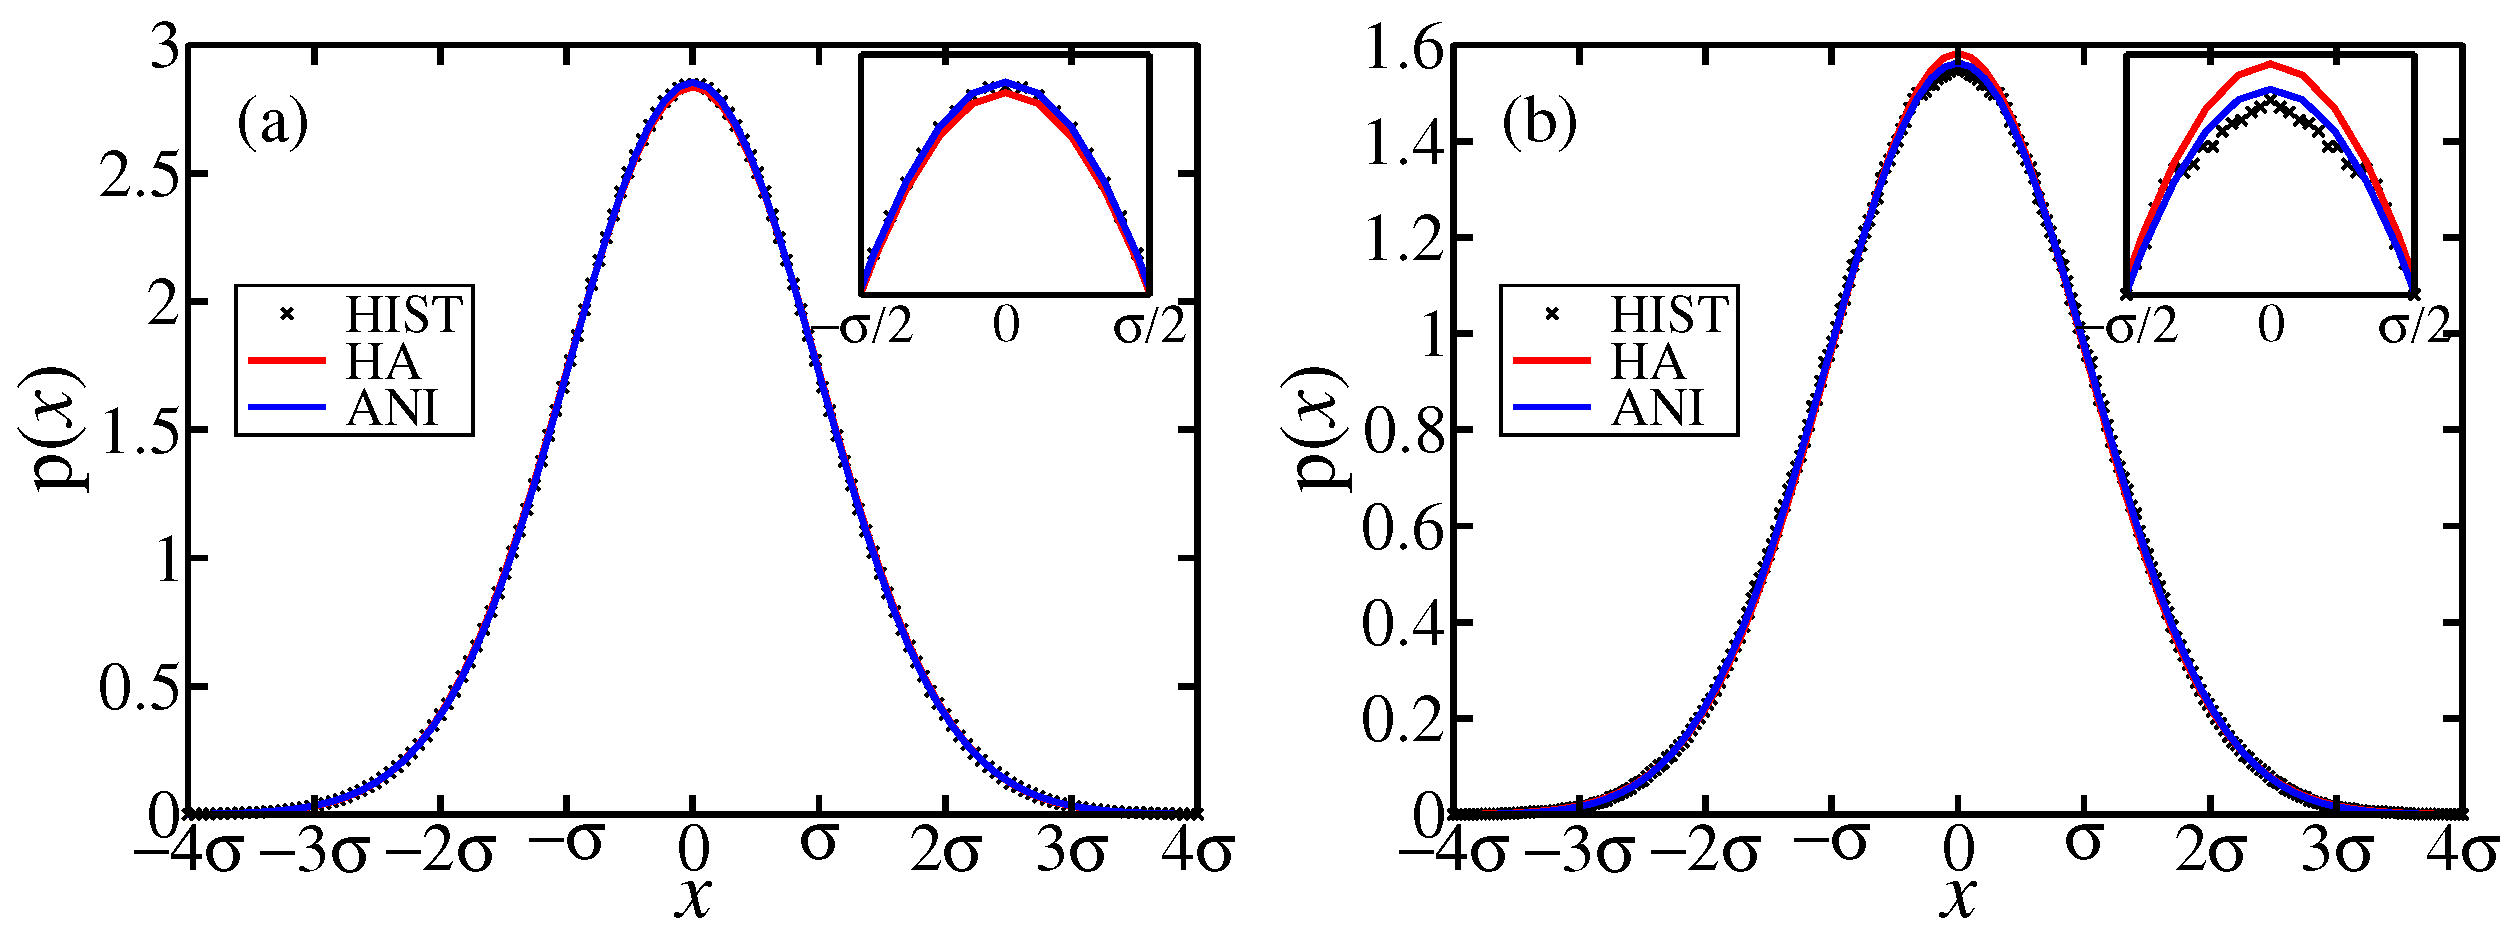
3.3.1. Anharmonicity
4. Conclusions
Author Contributions
Funding
Institutional Review Board Statement
Informed Consent Statement
Data Availability Statement
Acknowledgments
Conflicts of Interest
References
- Hoover, W.G.; Ree, F.H. Use of Computer Experiments to Locate the Melting Transition and Calculate the Entropy in the Solid Phase. J. Chem. Phys. 1967, 47, 4873. [Google Scholar] [CrossRef] [Scilit]
- Frenkel, D.; Laad, A.J.C. New Monte Carlo method to compute the free energy of arbitrary solids. Application to the fcc and hcp phases of hard spheres. J. Chem. Phys. 1984, 81, 3188. [Google Scholar] [CrossRef] [Scilit]
- Morris, J.R.; Ho, K.M. Calculating Accurate Free Energies of Solids Directly from Simulations. Phys. Rev. Lett. 1995, 74, 940–943. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Meirovitch, H.; Cheluvaraja, S.; White, R.P. Methods for calculating the entropy and free energy and their application to problems involving protein flexibility and ligand binding. Curr. Prot. Pept. Sci. 2009, 10, 229–243. [Google Scholar] [CrossRef] [Scilit]
- Widom, M. Entropy and diffuse scattering: Comparison of NbTiVZr and CrMoNbV. Met. Mater. Trans. A 2016, 47, 3306–3311. [Google Scholar] [CrossRef] [Scilit]
- Sutton, C.; Levchenko, S.V. First-Principles Atomistic Thermodynamics and Configurational Entropy. Front. Chem. 2020, 8, 757. [Google Scholar] [CrossRef] [Scilit]
- Nir, A.; Sela, E.; Beck, R.; Bar-Sinai, Y. Machine-learning iterative calculation of entropy for physical systems. Proc. Natl. Acad. Sci. USA 2020, 117, 30234–30240. [Google Scholar] [CrossRef] [Scilit]
- Nicholson, D.M.; Gao, C.Y.; McDonnell, M.T.; Sluss, C.C.; Keffer, D.J. Entropy Pair Functional Theory: Direct Entropy Evaluation Spanning Phase Transitions. Entropy 2021, 23, 234. [Google Scholar] [CrossRef] [Scilit]
- Sluss, C.C.; Pittman, J.; Nicholson, D.M.; Keffer, D.J. Exploration of Entropy Pair Functional Theory. Entropy 2022, 4, 603. [Google Scholar] [CrossRef] [Scilit]
- Hellman, O.; Abrikosov, I.; Simak, S. Lattice dynamics of anharmonic solids from first principles. Phys. Rev. B 2011, 84, 180301. [Google Scholar] [CrossRef] [Scilit]
- Hellman, O.; Steneteg, P.; Abrikosov, I.A.; Simak, S.I. Temperature dependent effective potential method for accurate free energy calculations of solids. Phys. Rev. B 2013, 87, 104111. [Google Scholar] [CrossRef] [Scilit]
- Frenkel, D.; Smit, B. Understanding Molecular Simulation: From Algorithms to Applications, 2nd ed.; Academic Press: Cambridge, MA, USA, 2002. [Google Scholar]
- Kastner, J.; Thiel, W. Bridging the gap between thermodynamic integration and umbrella sampling provides a novel analysis method: “Umbrella integration”. J. Chem. Phys. 2005, 123, 144104. [Google Scholar] [CrossRef] [Scilit] [PubMed]
- Grabowski, B.; Ikeda, Y.; Srinivasan, P.; Körmann, F.; Freysoldt, C.; Duff, A.I.; Shapeev, A.; Neugebauer, J. Ab initio vibrational free energies including anharmonicity for multicomponent alloys. NPJ Comp. Mater. 2019, 5, 80. [Google Scholar] [CrossRef] [Scilit]
- Fultz, B. Vibrational thermodynamics of materials. Prog. Mater. Sci. 2010, 55, 247–352. [Google Scholar] [CrossRef] [Scilit]
- Widom, M. Modeling the structure and thermodynamics of high entropy alloys. J. Mater. Res. 2018, 33, 2881–2898. [Google Scholar] [CrossRef] [Scilit]
- Ma, D.; Grabowski, B.; Körmann, F.; Neugebauer, J.; Raabe, D. Ab initio thermodynamics of the CoCrFeMnNi high entropy alloy: Importance of entropy contributions beyond the configurational one. Acta Mater. 2015, 100, 90–97. [Google Scholar] [CrossRef] [Scilit]
- Gao, M.C.; Widom, M. Information entropy of liquid metals. J. Phys. Chem. B 2018, 122, 3550–3555. [Google Scholar] [CrossRef] [Scilit]
- Widom, M.; Gao, M.C. First principles calculation of the entropy of liquid aluminum. Entropy 2019, 21, 131. [Google Scholar] [CrossRef] [Scilit]
- Huang, Y.; Gao, M.C.; Widom, M. ab initio free energies of liquid metal alloys: Application to the phase diagrams of Li-Na and K-Na. Phys. Rev. Mater. 2021, 6, 013802. [Google Scholar] [CrossRef] [Scilit]
- Allison, T. JANAF Thermochemical Tables, NIST Standard Reference Database 13. 1996. Available online: https://janaf.nist.gov/tables/Al-001.html (accessed on 23 April 2022).
- Allison, T. JANAF Thermochemical Tables, NIST Standard Reference Database 13. 1996. Available online: https://janaf.nist.gov/tables/Na-001.html (accessed on 23 April 2022).
- Nakashima, P.N. The Crystallography of Aluminum and Its Alloys. In Encyclopedia of Aluminum and Its Alloys; CRC Press: Boca Raton, FL, USA, 2019. [Google Scholar]
- Foust, O.J. Sodium-Nak Engineering Handbook; Gordon and Breach: New York, NY, USA, 1972. [Google Scholar]
- Rahman, A. Correlations in the Motion of Atoms in Liquid Argon. Phys. Rev. 1964, 136, 405–411. [Google Scholar] [CrossRef] [Scilit]
- Dickey, J.M.; Paskin, A. Computer Simulation of the Lattice Dynamics of Solids. Phys. Rev. 1969, 188, 1407–1418. [Google Scholar] [CrossRef] [Scilit]
- Lin, S.T.; Blanco, M.; Goddard, W.A. The two-phase model for calculating thermodynamic properties of liquids from molecular dynamics: Validation for the phase diagram of Lennard-Jones fluids. J. Chem. Phys. 2003, 119, 11792–11805. [Google Scholar] [CrossRef] [Scilit]
- Senkov, O.N.; Wilks, G.B.; Miracle, D.B.; Chuang, C.P.; Liaw, P.K. Refractory high-entropy alloys. Intermetallics 2010, 18, 1758–1765. [Google Scholar] [CrossRef] [Scilit]
- Widom, M.; Huhn, W.; Maiti, S.; Steurer, W. Hybrid Monte Carlo/molecular dynamics simulation of a refractory metal high entropy alloy. Met. Mater. Trans. A 2014, 45, 196–200. [Google Scholar] [CrossRef] [Scilit]
- Gibbs, J.W. Elementary Principles in Statistical Mechanics; Yale: New Haven, CT, USA, 1902. [Google Scholar]
- Shannon, C.E. A Mathematical Theory of Communication. Bell Syst. Tech. J. 1948, 27, 379–423. [Google Scholar] [CrossRef] [Scilit]
- Schultz, A.J.; Kofke, D.A. Comprehensive high-precision high-accuracy equation of state and coexistence properties for classical Lennard-Jones crystals and low-temperature fluid phases. J. Chem. Phys. 2018, 149, 204508. [Google Scholar] [CrossRef] [Scilit]
- Nezhad, S.Y.; Deiters, U.K. Estimation of the entropy of fluids with Monte Carlo computer simulation. Mol. Phys. 2016, 115, 1074–1085. [Google Scholar] [CrossRef] [Scilit]
- Kresse, G.; Furthmuller, J. Efficient iterative schemes for ab initio total-energy calculations using a plane-wave basis set. Phys. Rev. B 1996, 54, 11169–11186. [Google Scholar] [CrossRef] [Scilit]
- Kresse, G.; Joubert, D. From ultrasoft pseudopotentials to the projector augmented-wave method. Phys. Rev. B 1999, 59, 1758–1775. [Google Scholar] [CrossRef] [Scilit]
- Perdew, J.P.; Burke, K.; Ernzerhof, M. Generalized Gradient Approximation Made Simple. Phys. Rev. Lett. 1996, 77, 3865. [Google Scholar] [CrossRef] [Scilit]
- Togo, A.; Tanaka, I. First principles phonon calculations in materials science. Scr. Mater. 2015, 108, 1–5. [Google Scholar] [CrossRef] [Scilit]
- Togo, A.; Chaput, L.; Tanaka, I.; Hug, G. First-principles phonon calculations of thermal expansion in Ti3SiC2, Ti3AlC2, and Ti3GeC2. Phys. Rev. B 2010, 81, 174301. [Google Scholar] [CrossRef] [Scilit]
- Electronic Free Energy (Felect). Available online: http://alloy.phys.cmu.edu/Felect.html (accessed on 23 April 2022).
- Warren, B.E. X-ray Diffraction; Addison-Wesley: Reading, MA, USA, 1969. [Google Scholar]
- Jeong, I.K.; Heffner, R.H.; Graf, M.J.; Billinge, S.J.L. Lattice dynamics and correlated atomic motion from the atomic pair distribution function. Phys. Rev. B 2003, 67, 104301. [Google Scholar] [CrossRef] [Scilit]
- Gao, M.C.; Zhang, C.; Gao, P.; Zhang, F.; Ouyang, L.Z.; Widom, M.; Hawk, J.A. Thermodynamics of concentrated solid solution alloys. Curr. Opin. Solid State Mater. Sci. 2017, 21, 238–251. [Google Scholar] [CrossRef] [Scilit]
- Körmann, F.; Ikeda, Y.; Grabowski, B.; Sluiter, M.H.F. Phonon broadening in high entropy alloys. NPJ Comp. Mater. 2017, 3, 36. [Google Scholar] [CrossRef] [Scilit]
- Allison, T. JANAF Thermochemical Tables, NIST Standard Reference Database 13. 1996. Available online: https://janaf.nist.gov/tables/Mo-001.html (accessed on 23 April 2022).
- Allison, T. JANAF Thermochemical Tables, NIST Standard Reference Database 13. 1996. Available online: https://janaf.nist.gov/tables/Nb-001.html (accessed on 23 April 2022).
- Allison, T. JANAF Thermochemical Tables, NIST Standard Reference Database 13. 1996. Available online: https://janaf.nist.gov/tables/Ta-001.html (accessed on 23 April 2022).
- Allison, T. JANAF Thermochemical Tables, NIST Standard Reference Database 13. 1996. Available online: https://janaf.nist.gov/tables/W-001.html (accessed on 23 April 2022).
- Burgers, W. On the process of transition of the cubic-body-centered modification into the hexagonal-close-packed modification of zirconium. Physica 1934, 1, 561–586. [Google Scholar] [CrossRef] [Scilit]
- Feng, B.; Widom, M. Band structure theory of the bcc to hcp Burgers distortion. Phys. Rev. B 2018, 98, 174108. [Google Scholar] [CrossRef] [Scilit]
- van de Walle, A.; Hong, Q.; Kadkhodaei, S.; Sun, R. The free energy of mechanically unstable phases. Nat. Commun. 2015, 6, 7559. [Google Scholar] [CrossRef] [Scilit]
- Kadkhodaei, S.; Hong, Q.J.; van de Walle, A. Free energy calculation of mechanically unstable but dynamically stabilized bcc titanium. Phys. Rev. B 2017, 95, 064101. [Google Scholar] [CrossRef] [Scilit]
- Kadkhodaei, S.; van de Walle, A. Software tools for thermodynamic calculation of mechanically unstable phases from first-principles data. Comp. Phys. Commun. 2020, 246, 106712. [Google Scholar] [CrossRef] [Scilit]
- Mei, Z.G.; Shang, S.L.; Wang, Y.; Liu, Z.K. Density-functional study of the thermodynamic properties and the pressure–temperature phase diagram of Ti. Phys. Rev. B 2009, 80, 104116. [Google Scholar] [CrossRef] [Scilit]
- Sangiovanni, D.; Klarbring, J.; Smirnova, D.; Skripnyak, N.; Gambino, D.; Mrovec, M.; Simak, S.; Abrikosov, I. Superioniclike diffusion in an elemental crystal: Bcc Titanium. Phys. Rev. Lett. 2019, 123, 105501. [Google Scholar] [CrossRef] [Scilit]
- Allison, T. JANAF Thermochemical Tables, NIST Standard Reference Database 13. 1996. Available online: https://janaf.nist.gov/tables/Ti-001.html (accessed on 23 April 2022).
- Aurelio, G.; Fernández Guillermet, A.; Cuello, G.J.; Campo, J. Metastable phases in the Ti-V system: Part I. Neutron diffraction study and assessment of structural properties. Met. Mater. Trans. A 2002, 33, 1307–1317. [Google Scholar] [CrossRef] [Scilit]
- Spreadborough, J.; Christian, J.W. The Measurement of the Lattice Expansions and Debye Temperatures of Titanium and Silver by X-ray Methods. Proc. Phys. Soc. 1959, 74, 609–615. [Google Scholar] [CrossRef] [Scilit]
- Pawar, R.R.; Deshpande, V.T. The anisotropy of the thermal expansion of α-titanium. Acta Cryst. A 1968, 24, 316–317. [Google Scholar] [CrossRef] [Scilit]
- Roberts, W. Preferred orientation and anisotropy in titanium. J. Less-Common Met. 1962, 4, 345–361. [Google Scholar] [CrossRef] [Scilit]
- Allison, T. JANAF Thermochemical Tables, NIST Standard Reference Database 13. 1996. Available online: https://janaf.nist.gov/tables/Ti-002.html (accessed on 23 April 2022).
- Allison, T. JANAF Thermochemical Tables, NIST Standard Reference Database 13. 1996. Available online: https://janaf.nist.gov/tables/Ti-003.html (accessed on 23 April 2022).
- Petry, W.; Heiming, A.; Trampenau, J.; Alba, M.; Herzig, C.; Schober, H.R.; Vogl, G. Phonon dispersion of the bcc phase of group-IV metals. I, bcc titanium. Phys. Rev. B 1991, 43, 10933–10947. [Google Scholar] [CrossRef] [Scilit]
- Belonoshko, A.B.; Lukinov, T.; Fu, J.; Zhao, J.; Davis, S.; Simak, S.I. Stabilization of body-centred cubic iron under inner-core conditions. Nat. Geosci. 2017, 10, 312–316. [Google Scholar] [CrossRef] [Scilit]
- Perdew, J.P.; Zunger, A. Self-interaction correction to density functional approximations for many-electron systems. Phys. Rev. B 1981, 23, 5048–5079. [Google Scholar] [CrossRef] [Scilit]
- Sun, J.; Ruzsinszky, A.; Perdew, J.P. Strongly Constrained and Appropriately Normed Semilocal Density Functional. Phys. Rev. Lett. 2015, 115, 036402. [Google Scholar] [CrossRef] [Scilit]




| [Å] | [Å] | [Å] | [] | |
|---|---|---|---|---|
| FCC Al | 0.05933 | 0.005945 | 0.00003873 | 5.5450 |
| BCC Ti | 0.19061 | 0.06040 | −0.002597 | 9.4353 |
| [Å] | a | b | [] | |
| FCC Al | 0.01918 | −0.005718 | −0.001890 | −0.00017975 |
| BCC Ti | 0.06433 | 0.003210 | 0.01596 | −0.001227 |
Publisher’s Note: MDPI stays neutral with regard to jurisdictional claims in published maps and institutional affiliations. |
© 2022 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license (https://creativecommons.org/licenses/by/4.0/).
Share and Cite
Huang, Y.; Widom, M. Vibrational Entropy of Crystalline Solids from Covariance of Atomic Displacements. Entropy 2022, 24, 618. https://doi.org/10.3390/e24050618
Huang Y, Widom M. Vibrational Entropy of Crystalline Solids from Covariance of Atomic Displacements. Entropy. 2022; 24(5):618. https://doi.org/10.3390/e24050618
Chicago/Turabian StyleHuang, Yang, and Michael Widom. 2022. "Vibrational Entropy of Crystalline Solids from Covariance of Atomic Displacements" Entropy 24, no. 5: 618. https://doi.org/10.3390/e24050618
APA StyleHuang, Y., & Widom, M. (2022). Vibrational Entropy of Crystalline Solids from Covariance of Atomic Displacements. Entropy, 24(5), 618. https://doi.org/10.3390/e24050618



