


Abstract
A phytoalexin from papaya fruit has been synthesized in four steps; this procedure involved a Pummerer- type reaction.
Introduction
From papaya slices treated with copper salt or infected with Collectotrichum gloesporioides we isolated α-hydroxyacetosyringone as a phytoalexin [1,2]; recently, this compound was involved in plant-pathogen interactions, e.g. it is the major virulence gene activating factor and promotes high efficiency transformation of Arabidopsis thaliana explants by Agrobacterium tumefaciens [3,4].
Although its synthesis has been reported previously [5], we report here a four-steps synthesis of this compound
Results and Discussion
We attempted to synthesize α-hydroxyacetosyringone through a sequence of reactions including a Pummerer reaction. According to Russell and Becker [6,7], aromatic β-ketosulphoxides can be used to extend the carbon chain and also to add the desired new functional group. These β-ketosulphoxides are produced by reaction between DMSO and aldehydes or ketones in basic solutions and oxidation of the respective β-hydroxysulfoxide intermediate with MnO2 .
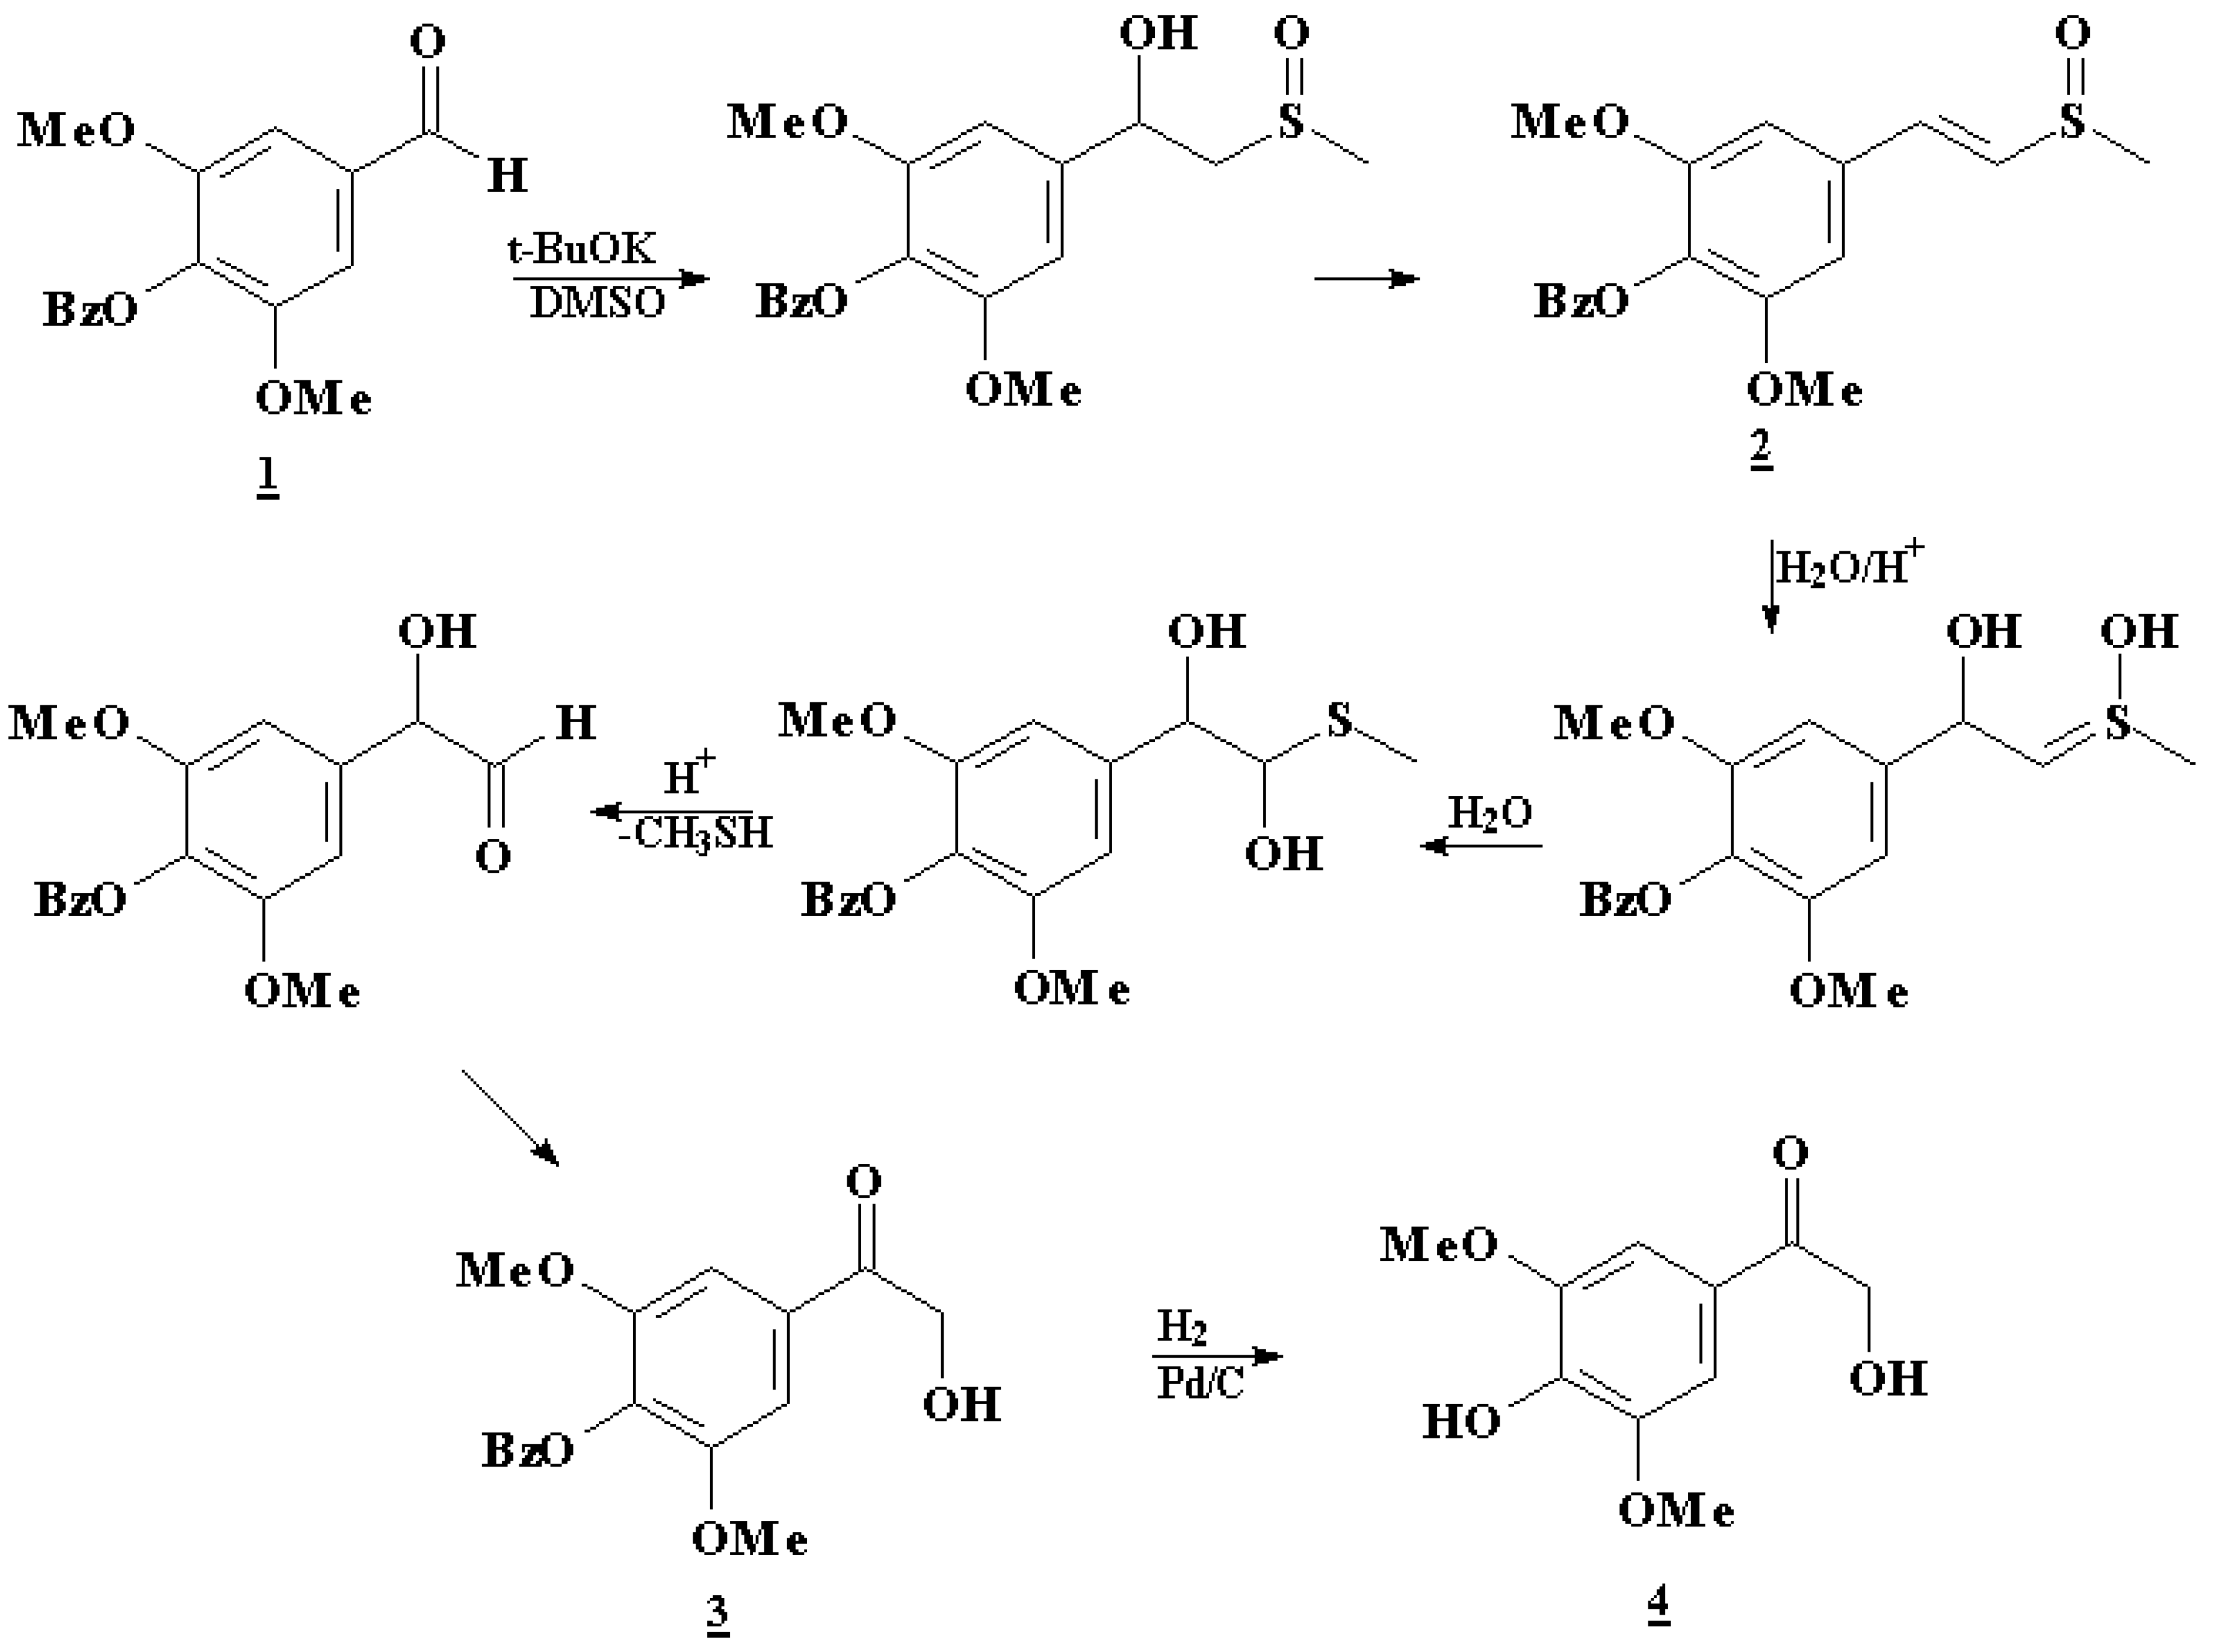
However, β-hydroxysulfoxides can yielded α,β-unsaturated sulfoxides under acidic conditions or upon neutralization of the condensation reaction. Moreover, other unexpected compounds have been isolated too, and their formation involved addition of water to α,β-unsaturated sulfoxide in an acid-catalyzed process [8]. Surprisingly, α-hydroxyacetosyringone was directly produced, via attack of two water molecules and thiomethanol elimination in a pH-dependent sequence of reactions.
Thus, when this coupling reaction was carried out between 4-O-benzyl-syringaldehyde and DMSO product 2 was obtained. This product can be transformed into 3 through a Pummerer-type reaction (Scheme 1); treatment of this substance with concentrated hydrochloric acid during 5 hours yielded a mixture of compounds, including compound 3. After hydrogenation of 3 on Pd/C the desired product 4 was obtained (<10% yield from 2). Its spectroscopic data were identical to those α-hydroxy acetosyringone obtained from natural sources.
Scheme 1.
Reactions involved in acetosyringone synthesis.
Experimental
General
NMR spectra were recorded with a Bruker AMX III (at 300 MHz for 1H and 75.0 MHz for 13C). All NMR spectra were taken in CDCl3; MS on a VG Micromass ZAB-2F at 70 eV; IR spectra were registered with a Perkin Elmer 1600 (FTIR).. TLC and column chromatographies were run using Merck silica gel and, unless otherwise specified, developed with n-hexane-ethyl acetate (9:1, v/v)
4-Benzyloxysyringaldehyde (1)
The commercial compound 3,5-dimethoxy-4-hydroxy-benzaldehyde (syringaldehyde) (91.09 mg, 0.5 mmol) in MeOH (5 mL) was treated with benzyl chloride (0.12 mL, 1.0 mmol) and stirred for 8hours at room temperature. Compound 1 was purified by silica gel column. HRMS: 272.1049. 1H NMR: 3.91 (6H, s, x 2-OCH3), 5.14 (2H, s, ArCH2O-), 7.12 (2H, s, H-3, H-5), 7.35 (3H, m, Ar), 7.47 (2H, d, J=6.1, Ar), 9.78 (1H, s, CHO). 13C NMR: 53.89 (q, - OCH3), 75.69 (t, -O-CH2), 107.31 (d, C-3, C-5), 129.08 (s, C-1), 143.01 (s, C-2,C-6), 154.63 (s, C-4), 191.83 (s, CHO).
3’,5’-Dimethoxy-4’-benzyloxyethenylmethylsulfoxide (2)
A solution of t-BuOK (179.55 mg, 1.6 mmol) in DMSO (5.0 mL) was added to 4-O-benzyl-syringaldehyde (272.30 mg, 1.0 mmol) 1, in DMSO (3.0 mL) under argon and stirred overnight. After extraction with ethyl acetate and purification by column chromatography, compound 2 was obtained (199.45 mg, 60%). TLC: Rf = 0.35; mp 102-104 oC. HRMS: 332.1082. 1H NMR 300 MHz (δ ppm CDCl3) 2.70 (3H, s, S–CH3), 3.83 (6H, s, –OMe), 5.02 (2H, s, ArCH2O-), 6.67 (2H, s, H-2’ and H-6’), 6.80 (1H, d, 14.0 Hz, =CH–S), 7.14 (1H, d, 14.0 Hz, Ar–CH=), 7.28-7.36 (3H, m, Ar), 7.46 (2H, dd, 2.0 and 7.0 Hz, Ar); 13C NMR 75.46 MHz: 41.01 (q, S-CH3), 56.10 (q, O-CH3), 75.06 (t, -ArCH2O-), 104.70 (d, C-2, C-5), 129.14 (d, -S-C=CH), 128.15 (d, Bz), 128.44 (s, C-1), 136.51 (d, S-CH=C), 137.41 (s, C-1’), 138.31 (s, C-4), 153.71 (s, C-3, C-5).
4’-O-benzyl-α-hydroxyacetosyringone (3)
The sulfoxide 2 (166.21 mg, 0.5 mmol) was added to 10% HCl (10 mL) and heated to reflux for five hours. The crude reaction mixture was purified by TLC (silicagel; n-hexane-ethyl acetate 6:1, v/v) and product 3 was thus obtained as a yellow oil (15.11 mg, 10%). HRMS: 302.1154. 1H NMR : 3.68 (6H, s, x 2 OCH3), 5.07 (2H, s, -COCH2OH), 5.08 (2H, s, Ar-OCH2-), 6.48 (1H, s, -OH), 6.90 (2H, s, H-2’ and H-6’), 7.30 (5H, m, benzyl); 13C NMR: 56.49 (q, x 2 OCH3), 75.39 (t, x 2CH2), 107.40 (d, C-2’ and C-5’ ), 128.80 (d, Bz), 132. 90 (s, C-1’), 134.80 (s, C-1’’), 138.87 (s, C-4’), 153.45 (s, C-3’, C-5’), 192.15 (s, CO).
α-Hydroxyacetosyringone (4)
Compound 3 (15.11 mg, 0.05 mmol) in MeOH (5 mL) was hydrogenated at room temperature for 5 hours over Pd/C and after workup compound 4 was recovered (10 mg, yield 95%). Its spectroscopical and physical properties were compared to an authentic sample obtained from papaya slices. mp. 145 º, Rf = 0.28 (n-hexane-ethyl acetate 2:3). HRMS: 212.0685. 1H NMR : 3.25 (H, s, 2-OH), 3.89 (6H, s, 3’,5’-OCH3), 4.93 (2 H, d, J=1.9, -COCH2OH), 6.34 (H, t, J=1.9, 4-OH), 7.28 (2H, s. H-2’ and H-6); 13C NMR: 54.95 (q, x 2 OCH3), 66.3 (t, C-2), 106.2 (d, C-2), 125.1 (s, C-1), 141.3 (s, C-4), 146.6(s, C-3, C-5), 196.4 (s, C-1).
Acknowledgments
Author thanks to COLCIENCIAS (Colombia) and Universidad de Antioquia for financial support.
References
- Echeverri, F.; Torres, F.; Quiñones, W.; Cardona, G.; Archbold, R.; Roldán, J.; Gutierrez, J.; Hassane, E. L. Phytochemistry 1996, 44, 255–256.
- Echeverri, F.; Torres, F.; Quiñones, W.; Cardona, G.; Archbold, R.; Roldán, J.; Brito, I.; Gutierrez, J.; Lahlou, E-H. “Memorias III Simposio Internacional de Química de Productos Naturales y sus Aplicaciones”; Sociedad Chilena de Química-U. de Chile, Santiago y Catolica de Chile: Punta de Tralca (Chile), 1996 December 4-7; pp. 125–126. [Google Scholar]
- Song, Y. N.; Shibuya, M.; Ebizuka, Y.; Sankawa, U. Chem. Pharm. Bull. 1990, 38, 2063–2065.
- Sheikholeslam, S. N.; Weeks, D.B. Plant Mol. Biol. 1987, 8, 291–298. [PubMed]
- Luis, J. G.; San Andres, L. J. Chem. Res. (S) 1999, 220–221. [CrossRef]
- Russell, G. A.; Becker, H. D. J. Am. Chem. Soc. 1963, 85, 3406–10.
- Becker, H. D.; Mikol, G.; Russell, G. J. Am. Chem. Soc. 1963, 85, 3410–3414.
- Russell, G.; Mikol, G. J. Am. Chem. Soc. 1966, 88, 5498–5504.
- Sample Availability: Samples are available from the authors.

© 2000 by MDPI (http://www.mdpi.org).