Journal Description
Viruses
Viruses
is a peer-reviewed, open access journal of virology, published monthly online by MDPI. The Spanish Society for Virology (SEV), Canadian Society for Virology (CSV), Italian Society for Virology (SIV-ISV), Australasian Virology Society (AVS), Brazilian Society for Virology (BSV) and Global Virus Network (GVN) are affiliated with Viruses and their members receive a discount on the article processing charges.
- Open Access— free for readers, with article processing charges (APC) paid by authors or their institutions.
- High Visibility: indexed within Scopus, SCIE (Web of Science), PubMed, MEDLINE, PMC, Embase, PubAg, and other databases.
- Journal Rank: JCR - Q2 (Virology) / CiteScore - Q1 (Infectious Diseases)
- Rapid Publication: manuscripts are peer-reviewed and a first decision is provided to authors approximately 15.5 days after submission; acceptance to publication is undertaken in 2.9 days (median values for papers published in this journal in the first half of 2026).
- Recognition of Reviewers: Reviewers whose reports are timely and of high quality receive an APC discount voucher for a future publication in an MDPI journal. Become a reviewer.
- Journal Cluster of Microbiology: Acta Microbiologica Hellenica, Applied Microbiology, Bacteria, Journal of Fungi, Microorganisms, Microbiology Research, Pathogens, Viruses, Fermentation and Germs.
Impact Factor:
3.8 (2025);
5-Year Impact Factor:
3.8 (2025)
Latest Articles
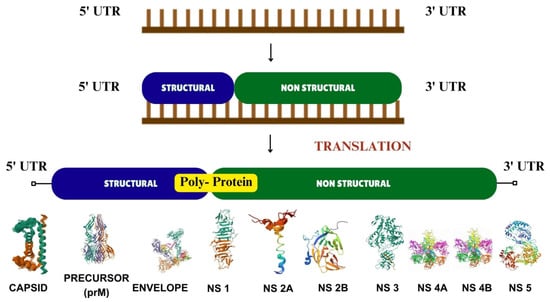
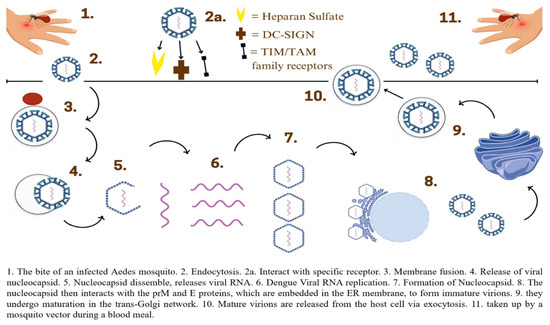
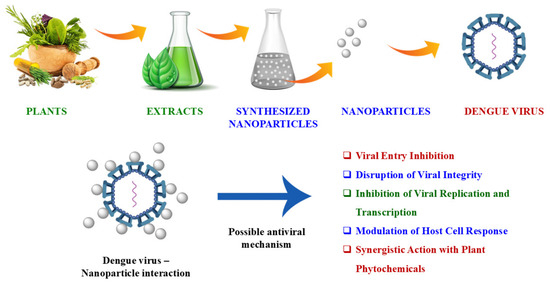
Targeting Dengue Viral Proteins with Eco-Friendly Nanotechnology: A Promising Path for Antiviral Research
Viruses 2026, 18(9), 995; https://doi.org/10.3390/v18090995 - 9 Sep 2026
Abstract
Dengue virus (DENV), a mosquito-borne pathogen belonging to the Flaviviridae family, continues to be a major global public health concern, causing millions of infections annually across tropical and subtropical regions. The virus encodes both structural and non-structural proteins that play critical roles in
[...] Read more.
Dengue virus (DENV), a mosquito-borne pathogen belonging to the Flaviviridae family, continues to be a major global public health concern, causing millions of infections annually across tropical and subtropical regions. The virus encodes both structural and non-structural proteins that play critical roles in its replication, assembly, and immune evasion. Despite substantial research progress, there is still no specific antiviral drug available, and current treatment remains primarily supportive. This review highlights the significance of DENV structural proteins (C, prM/M, and E) and non-structural proteins (NS1–NS5) as potential molecular targets for antiviral interventions. Green-synthesized silver, gold, and iron oxide nanoparticles have demonstrated strong inhibitory activity against DENV by targeting key viral proteins such as the envelope (E), NS3 protease/helicase, and NS5 RNA-dependent RNA polymerase. These nanoparticles interfere with viral entry, replication, and protein synthesis while exhibiting high biocompatibility and minimal cytotoxicity. Collectively, the findings discussed in this review underscore the promise of eco-friendly nanotechnology as a sustainable and effective platform for developing next-generation antiviral therapies against dengue virus.
Full article
(This article belongs to the Special Issue Natural, Semisynthetic, and Synthetic Antiviral Drugs: Combating Emerging and Re-Emerging Viral Threats)
►
Show Figures
Open AccessReview
Peripheral Blood Mononuclear Cell Transcriptomics in Porcine Reproductive and Respiratory Syndrome: A Window into Innate Immune Resistance and Tolerance to Viral Disease
by
Md Aminul Islam, Christiane Neuhoff, Maren Julia Pröll, Christine Große-Brinkhaus, Sharmin Aqter Rony, Ernst Tholen, Karl Schellander and Muhammad Jasim Uddin
Viruses 2026, 18(9), 994; https://doi.org/10.3390/v18090994 - 9 Sep 2026
Abstract
Peripheral blood mononuclear cells (PBMCs) are the most immunologically active and readily accessible fraction of whole blood, and they initiate host immune responses following infection and vaccination. Since PBMC transcriptomes capture diverse host–pathogen interactions, they offer a promising tool to uncover the genetic
[...] Read more.
Peripheral blood mononuclear cells (PBMCs) are the most immunologically active and readily accessible fraction of whole blood, and they initiate host immune responses following infection and vaccination. Since PBMC transcriptomes capture diverse host–pathogen interactions, they offer a promising tool to uncover the genetic drivers of viral resistance and tolerance. As a detailed case study of this principle, we examine porcine reproductive and respiratory syndrome (PRRS), which remains one of the most economically important viral diseases of swine worldwide. Following PRRS virus (PRRSV) exposure, pigs rely on two complementary defense strategies: resistance, the capacity to limit viral replication, and tolerance, the capacity to sustain performance despite infection. Since resistance and tolerance phenotypes are difficult to measure directly through experimental challenge, indirect immune-trait measurements collected after vaccination offer a practical alternative. Most transcriptomic studies of the host response to PRRSV have focused on respiratory tissues, reflecting the virus’s tropism for pulmonary macrophages. However, intramuscularly delivered modified-live PRRSV vaccine reaches the bloodstream, bypassing the lung, so PBMCs, as the frontline defense system, mount the earliest measurable innate response. This review synthesizes the current literature on PBMC transcriptome models for deciphering innate resistance and tolerance to viral disease, using PRRS as our principal worked example; presents our own approach to profiling PBMCs after PRRSV vaccination; and outlines how the field has advanced since the original candidate-gene and QTL studies of the 2010s, including the recent FDA approval of the first CD163 gene-edited PRRSV-resistant pig line, and the emergence of single-cell and multi-tissue PBMC atlases, before considering how the same PBMC-based approach could extend to other host–virus interactions.
Full article
(This article belongs to the Section Animal Viruses)
►▼
Show Figures

Figure 1
Open AccessReview
SARS-CoV-2 Drug-Resistant Mutations in Non-Structural Proteins
by
Madison Shaw and Anthony R. Fehr
Viruses 2026, 18(9), 993; https://doi.org/10.3390/v18090993 - 9 Sep 2026
Abstract
Various antiviral drug therapies have been developed for the treatment of coronaviruses, particularly SARS-CoV-2. However, the emergence of drug-resistant mutations is detrimental to clinical efficacy and global public health. Antiviral usage exerts selective pressures on viruses that manufacture an environment for resistant strains
[...] Read more.
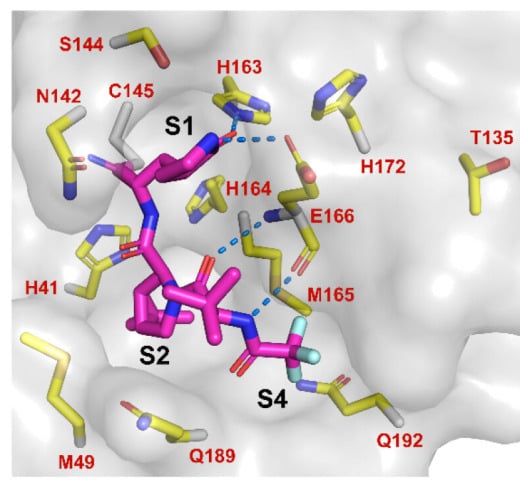
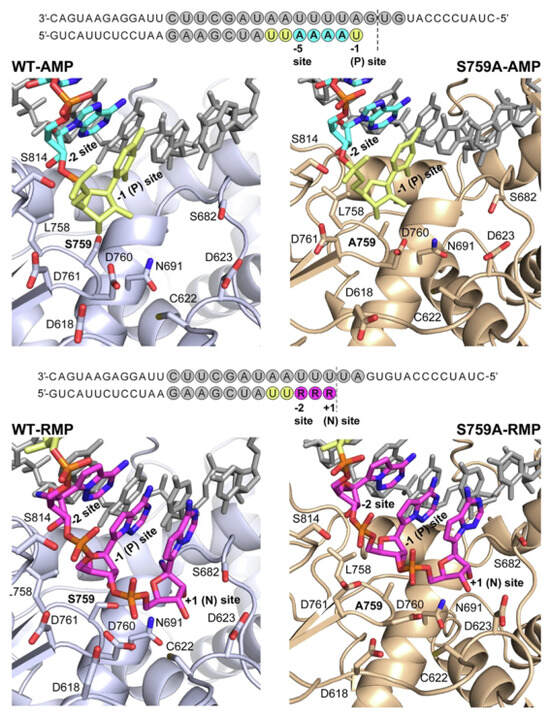
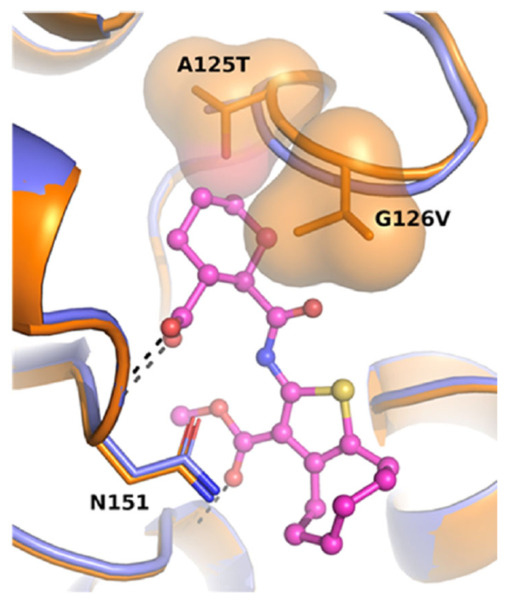
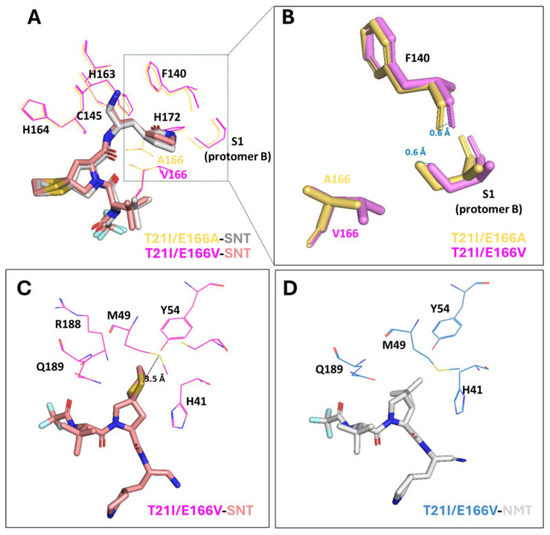
Various antiviral drug therapies have been developed for the treatment of coronaviruses, particularly SARS-CoV-2. However, the emergence of drug-resistant mutations is detrimental to clinical efficacy and global public health. Antiviral usage exerts selective pressures on viruses that manufacture an environment for resistant strains to emerge, in some cases at a fitness cost. However, the appearance of a compensatory mutation can restore or improve viral fitness, allowing the strain to persist and spread in a population. Here we evaluate the drug resistance mechanisms of multiple SARS-CoV-2 non-structural proteins, including the main protease (Mpro) and the RNA-dependent RNA polymerase (RdRP), which drive polyprotein processing and viral RNA replication, as well as PLPro, EndoU, and Mac1, which contribute to viral replication and counter host innate immune responses. We also discuss several methods that could be used to avoid drug resistance in the future. By integrating the understanding of molecular mechanisms of antiviral treatment with surveillance of resistance-associated mutations and new drug therapies, appropriate clinical approaches can be developed to reduce the impact of drug resistance.
Full article
(This article belongs to the Special Issue Coronaviruses Pathogenesis, Immunity, and Antivirals (2nd Edition))
►▼
Show Figures

Figure 1
Open AccessArticle
Etiologies and Diagnostic Yield of Bone Marrow Evaluation in Adults Living with HIV in Venezuela: A Cross-Sectional Study
by
Lily M. Soto-Avila, Higinio Fernández-Sánchez and Alfonso J. Rodriguez-Morales
Viruses 2026, 18(9), 992; https://doi.org/10.3390/v18090992 - 9 Sep 2026
Abstract
Background: Bone marrow abnormalities in people living with HIV may reflect infectious, neoplastic, or inflammatory processes, particularly in advanced disease presenting with fever of unknown origin or unexplained cytopenias. In Latin America, contemporary data on the causes and diagnostic contribution of bone marrow
[...] Read more.
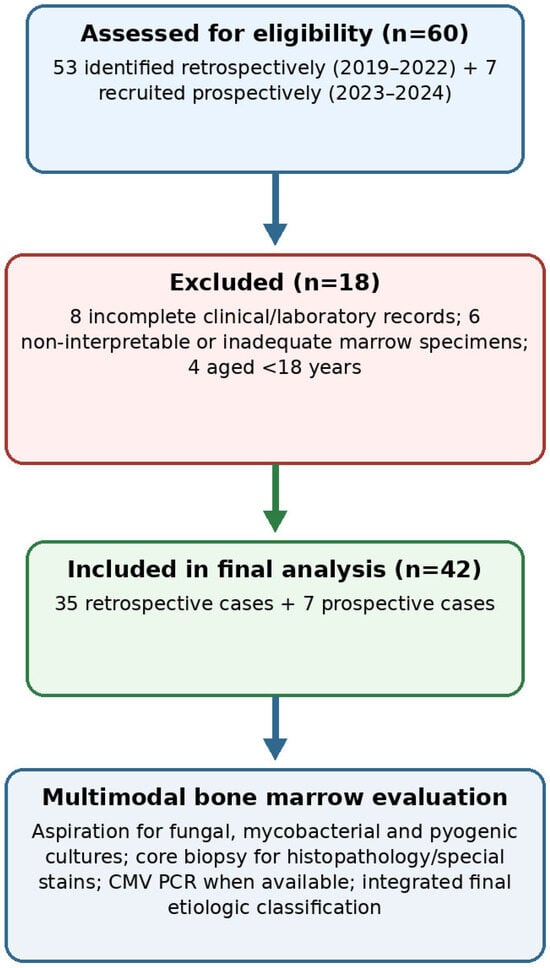
Background: Bone marrow abnormalities in people living with HIV may reflect infectious, neoplastic, or inflammatory processes, particularly in advanced disease presenting with fever of unknown origin or unexplained cytopenias. In Latin America, contemporary data on the causes and diagnostic contribution of bone marrow evaluation remain limited, especially in resource-constrained settings. Aim: To characterize the etiologies, histopathological patterns, and diagnostic contribution of bone marrow evaluation in adults living with HIV at a tertiary referral center in Venezuela. Methods: We conducted a cross-sectional study with retrospective and prospective case ascertainment among adults with confirmed HIV infection who underwent bone marrow aspiration, biopsy, or both for suspected infiltrative disease between January 2019 and April 2024. We analyzed clinical, laboratory, histopathological, microbiological, and molecular data at the index bone marrow evaluation. Test-specific denominators are reported because diagnostic investigations were not uniformly available. Results: Forty-two patients were included (35 retrospectively and 7 prospectively); 59.5% were male and the median age was 39 years (IQR: 31.75–50.0). At presentation, 67% were newly diagnosed with HIV and 79% were ART-naive; the median CD4+ T-cell count was 98.5 cells/mm3. Histopathology was evaluable in 37/42 patients; 31/37 (83.8%) showed findings compatible with infectious involvement and 6/37 (16.2%) showed malignancy. Histoplasma capsulatum was isolated in 17/35 fungal cultures (48.6%), Mycobacterium tuberculosis in 13/33 mycobacterial cultures (39.4%), and CMV PCR on bone marrow aspirate was positive in 2/3 patients tested. The final etiologic classification was infectious in 36/42 patients (85.7%) and neoplastic in 6/42 (14.3%). Conclusions: In this highly selected cohort of adults with advanced HIV disease, infectious etiologies-particularly histoplasmosis and tuberculosis-predominated. Bone marrow aspiration and biopsy, interpreted together with microbiological and molecular testing, provided clinically relevant diagnostic information in patients with prolonged fever, cytopenias, or otherwise inconclusive investigations. The findings support earlier HIV diagnosis and ART initiation and improved access to fungal, mycobacterial, and molecular diagnostics in resource-limited settings.
Full article
(This article belongs to the Special Issue HIV and HTLV Infections and Coinfections (2nd Edition))
►▼
Show Figures

Figure 1
Open AccessReview
Molecular Mechanisms of Zika Virus Entry into Host Cells
by
Haolong Cong, Xin Zhao, Wenhui Li, Rong Lei, Wenjun Zhao and Xiaodong Han
Viruses 2026, 18(9), 991; https://doi.org/10.3390/v18090991 - 9 Sep 2026
Abstract
The World Health Organization classifies mosquito-borne viruses as a major global public health threat with pandemic risk and urges all nations to strengthen pandemic preparedness. As a representative mosquito-borne flavivirus, ZIKV bears prominent pandemic capacity. Its distinctive neurotropic property and capability of vertical
[...] Read more.
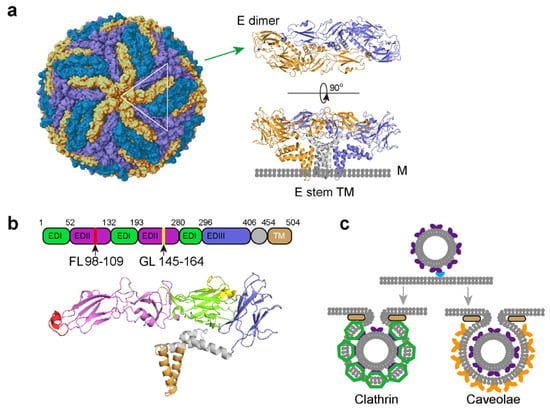
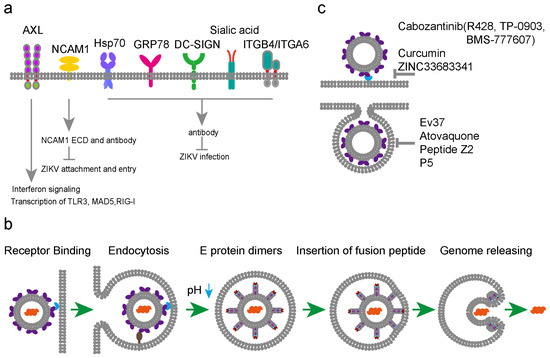
The World Health Organization classifies mosquito-borne viruses as a major global public health threat with pandemic risk and urges all nations to strengthen pandemic preparedness. As a representative mosquito-borne flavivirus, ZIKV bears prominent pandemic capacity. Its distinctive neurotropic property and capability of vertical transmission create unique pathogenic hazards and transmission risks. Owing to the absence of approved vaccines and targeted antiviral therapeutics, the prevention and management of ZIKV outbreaks remain extremely challenging. Viral entry into host cells marks the very first step of productive ZIKV infection, and mechanistic investigations into this entry process lay an essential theoretical foundation for developing small-molecule agents that block ZIKV cellular entry. This review summarizes recent advances concerning ZIKV host receptor usage, membrane fusion cascades, and entry-targeted inhibitor development; systematically discusses existing inconsistencies and core controversies in cell type-dependent entry pathways, functionally redundant host receptor usage, especially the disputed AXL receptor, and unresolved molecular details of membrane fusion resulting from diverse experimental models and technical limitations; and proposes targeted future research directions to resolve these bottlenecks and clarify the context-dependent ZIKV entry mechanisms.
Full article
(This article belongs to the Special Issue Preparation for the Next Potential Pandemic—Chikungunya, Dengue, Zika and Other Viruses 2026)
►▼
Show Figures

Figure 1
Open AccessEditorial
Special Issue, “Viruses and Eye Diseases: Emerging Mechanisms, Host–Virus Interactions, and Future Directions”
by
Deepak Shukla
Viruses 2026, 18(9), 990; https://doi.org/10.3390/v18090990 - 9 Sep 2026
Abstract
Vision-threatening viral infections remain a major cause of ocular morbidity and blindness worldwide despite substantial advances in antiviral therapy, molecular virology, and ocular immunology [...]
Full article
(This article belongs to the Special Issue Viruses and Eye Diseases)
Open AccessArticle
Identification of Serum Antibodies Cross-Reactive with Pathogenic Betacoronavirus Amongst Rural Communities Within the Forested Region of the Republic of Guinea
by
Tom R. W. Tipton, Joseph A. Bore, Joseph Timothy, Stephanie Longet, Stephen M. Laidlaw, Grace Hood, Craig Thompson, Jack Mellors, Ifono Kekoura, Millimon S. Lucien, Beatrice K. Koivogui, Kpade Zeze, David Matthews, Andrew D. Davidson, Alex Mentzer, Donal Skelly, Paul Klenerman, Julian Hiscox, N’Faly Magassouba, Kimberley Fornace and Miles W. Carrolladd
Show full author list
remove
Hide full author list
Viruses 2026, 18(9), 989; https://doi.org/10.3390/v18090989 - 8 Sep 2026
Abstract
Objectives: The SARS-CoV-2 pandemic caused major morbidity, mortality, and economic disruption, highlighting the need to better understand zoonotic spillover risks posed by coronaviruses. We investigated whether archived human sera from forested Guinea contained evidence of prior exposure to coronaviruses antigenically related to
[...] Read more.
Objectives: The SARS-CoV-2 pandemic caused major morbidity, mortality, and economic disruption, highlighting the need to better understand zoonotic spillover risks posed by coronaviruses. We investigated whether archived human sera from forested Guinea contained evidence of prior exposure to coronaviruses antigenically related to known pathogenic human coronaviruses. Methods: Archived sera collected in 2017–2018 from wildlife hunter communities in Guinea were analysed using serological assays against spike glycoproteins from SARS-CoV, SARS-CoV-2, MERS-CoV, and seasonal human coronaviruses. Binding responses to receptor binding domains were also assessed, together with spatial and subgroup analyses. Results: Pre-pandemic sera showed IgG cross-reactivity to SARS-CoV, SARS-CoV-2, and MERS-CoV spike antigens. A distinct subgroup demonstrated strong binding to both SARS-CoV and MERS-CoV receptor binding domains. Conclusions: These findings highlight the utility of sero-epidemiology in identifying potential zoonotic spillover. However, further investigation is needed to identify these viruses and determine their homology to known pathogenic coronaviruses.
Full article
(This article belongs to the Section Human Virology and Viral Diseases)
Open AccessArticle
Parvovirus B19 and Cellular Transcriptome Dynamics in UT7/EpoS1 Cells
by
Niccolò Guglietta, Federica Bichicchi, Ilaria Gasperini, Elisabetta Manaresi and Giorgio Gallinella
Viruses 2026, 18(9), 988; https://doi.org/10.3390/v18090988 - 8 Sep 2026
Abstract
Parvovirus B19 (B19V) is a human ssDNA virus with ample pathogenic potential, characterized by a selective tropism for erythroid progenitor cells (EPCs) in the bone marrow. In vitro, in addition to EPCs, UT7/EpoS1 cells are widely used as a model cell system, permissive
[...] Read more.
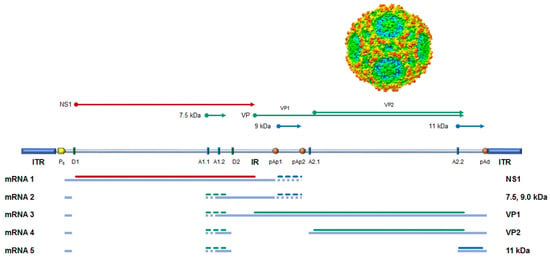
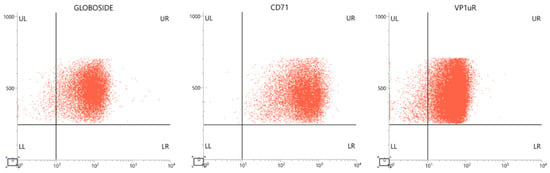
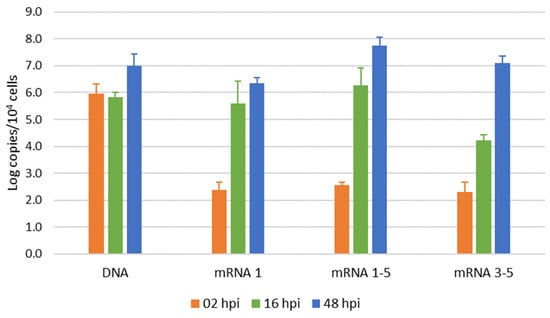
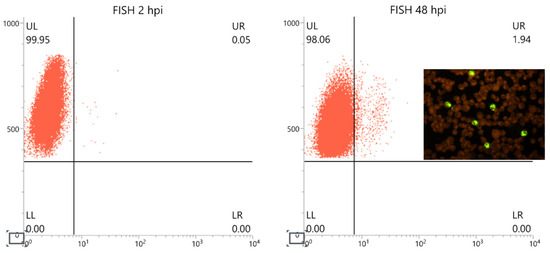
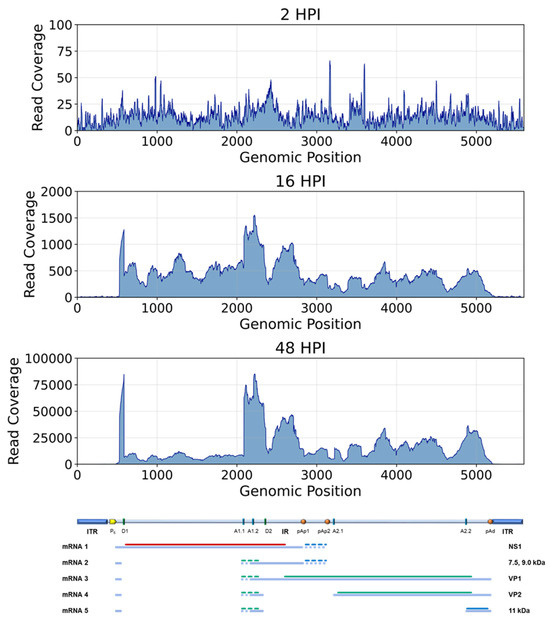
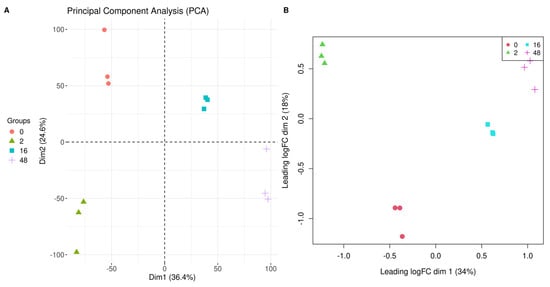
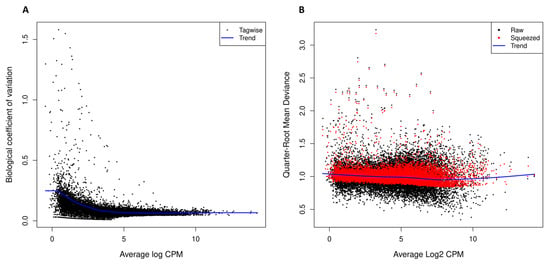
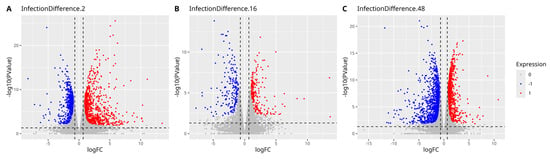
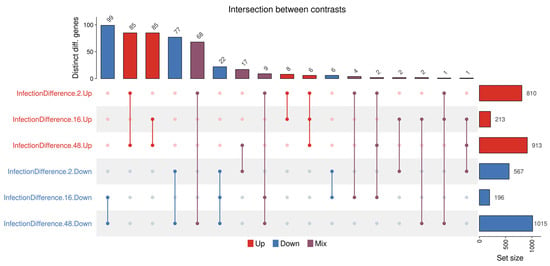
Parvovirus B19 (B19V) is a human ssDNA virus with ample pathogenic potential, characterized by a selective tropism for erythroid progenitor cells (EPCs) in the bone marrow. In vitro, in addition to EPCs, UT7/EpoS1 cells are widely used as a model cell system, permissive to viral replication, although in a restrictive pattern. In our work, we applied mRNA high-throughput sequencing technology (HTS) and a dedicated bioinformatic pipeline to investigate both viral and cellular expression profiles in the course of B19V infection of UT7/EpoS1 cells. Mapping of the viral transcriptome detailed the differential expression pattern across early and late time points in the course of infection, at 2, 16 and 48 h post-infection (hpi). Analysis of the cellular transcriptome indicated that downregulation of genes involved in the immune/cytokine/interleukin response was prominent from earlier time points throughout the time course of infection. Upregulation of genes involved in cell stress response was found at 2 hpi, and genes involved in cell cycle regulation were affected mainly at 16 hpi and 48 hpi. A comparative analysis was performed with EPCs, showing similarity in their viral expression profile but substantial divergence in the virus-induced dysregulation of the cellular transcription pattern. This dual-transcriptome analysis of infected UT7/EpoS1 cells and comparison with EPCs provides groundwork for future research aimed at providing a better definition of the pathogenic mechanisms of B19V.
Full article
(This article belongs to the Collection Parvoviridae)
►▼
Show Figures

Figure 1
Open AccessArticle
Public Information Demand During the 2026 Bundibugyo Virus Outbreak: A Dual Infodemiology Analysis Across Six Ebola Virus Disease Outbreaks
by
Hakkı Öztürk and Berrak Itır Aylı
Viruses 2026, 18(9), 987; https://doi.org/10.3390/v18090987 - 8 Sep 2026
Abstract
The 2026 Bundibugyo virus disease outbreak in the Democratic Republic of the Congo and Uganda, the first Ebola-related PHEIC caused by a species lacking a licensed vaccine, has been accompanied by an information crisis marked by conspiracy-driven violence against response teams and destruction
[...] Read more.
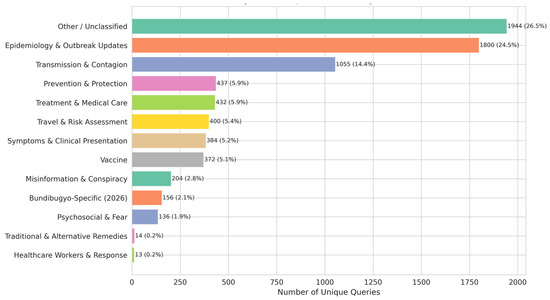
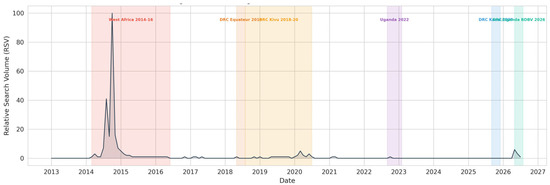
The 2026 Bundibugyo virus disease outbreak in the Democratic Republic of the Congo and Uganda, the first Ebola-related PHEIC caused by a species lacking a licensed vaccine, has been accompanied by an information crisis marked by conspiracy-driven violence against response teams and destruction of treatment centres. Understanding what populations actually seek to know during such events is essential for designing effective communication strategies, yet the content of public information demand during Ebola outbreaks has not been systematically characterised. We combined Google Autocomplete query extraction (7347 unique queries from 83 seed terms in English and French) with Google Trends temporal analysis across six Ebola outbreaks (2014–2026). The autocomplete suggestion landscape was dominated by risk quantification and transmission concerns. Google Trends temporal analysis showed moderate to strong co-movement between misinformation-related search terms and overall Ebola search interest (Spearman ρ = 0.359–0.697, nominal p < 0.001). Substantial differences emerged between the English-language and French-language suggestion sets: French-language suggestions contained six times more misinformation and conspiracy content (4.9% vs. 0.8%) and nearly nine times more fear-related content (3.5% vs. 0.4%) than English-language suggestions. These findings demonstrate that infodemic management strategies designed in Anglophone contexts are inadequate for the populations most at risk and should inform linguistically adapted interventions for the ongoing response.
Full article
(This article belongs to the Special Issue The 2026 Central Africa (Bundibugyo) Ebola Virus Outbreak—and Lessons Learned from Previous Ebola Virus Episodes)
►▼
Show Figures

Figure 1
Open AccessArticle
Late Clinical Presentation Outweighs Viral Characteristics in Determining Immune Depletion at HIV Diagnosis: A Five-Year Retrospective Study
by
Silvere D. Zaongo, Mei Han and Yaokai Chen
Viruses 2026, 18(9), 986; https://doi.org/10.3390/v18090986 - 8 Sep 2026
Abstract
Background: Late presentation and poor immunovirological outcomes persist among people living with HIV (PLWH). The differential impact of HIV-1 subtypes on baseline disease presentation in antiretroviral therapy (ART)-naïve individuals remains incompletely characterized, particularly in regions dominated by unique recombinant forms. Methods: We conducted
[...] Read more.
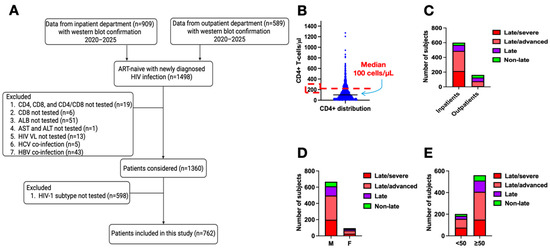
Background: Late presentation and poor immunovirological outcomes persist among people living with HIV (PLWH). The differential impact of HIV-1 subtypes on baseline disease presentation in antiretroviral therapy (ART)-naïve individuals remains incompletely characterized, particularly in regions dominated by unique recombinant forms. Methods: We conducted a retrospective cross-sectional study involving ART-naïve newly diagnosed PLWH at our hospital from January 2020 to October 2025. Immunological (CD4+, CD8+ T-cell counts, and CD4+/CD8+ ratios), biochemical [albumin (ALB), alanine aminotransferase (ALT), aspartate aminotransferase (AST)], virological (HIV-1 subtype, genotypic drug resistance), and clinical (admission type, age) parameters were analyzed. Multivariate linear regression identified factors independently associated with CD4+ T-cell count, CD4+/CD8+ ratio, and ALB level. Results: The study population comprised 762 ART-naïve newly diagnosed PLWH who were predominantly male (87.5%), older (median age 59), and presented with advanced immunosuppression (median CD4+ count 100 cells/µL). CRF07_BC was the dominant subtype (57.4%). Significant subtype-specific differences were observed: CRF01_AE infection was associated with the lowest CD4+ counts (median 54 cells/µL, p = 0.0004) and CD4+/CD8+ ratios (p = 0.0005). Inpatient status, indicating more severe presentation, was independently associated with significantly lower CD4+ counts (β = −36.91, p < 0.0001) and ALB levels (β = −7.358, p < 0.0001). Contrary to typical immune aging, patients aged ≥50 years had higher CD4+ counts at diagnosis (105.5 vs. 74 cells/µL, p = 0.0039). Genotypic drug resistance (22.2% prevalence) showed no independent association with baseline immunological status. Conclusions: Our findings reveal a distinct profile of late HIV presentation in a contemporary Chinese cohort, characterized by advanced age and the predominance of CRF07_BC. The independent association of inpatient status, rather than HIV-1 subtype or drug resistance, with severe immunosuppression and hypoalbuminemia highlights clinical presentation as the paramount indicator of baseline disease severity. The unexpected immunological advantage in older adults warrants further investigation.
Full article
(This article belongs to the Special Issue Epidemiology and Prevention of HIV/AIDS)
►▼
Show Figures

Figure 1
Open AccessReview
Molecular Mechanisms Between Canine Parvovirus 2 and Host Interactions: From Viruses to Innate Immune Signaling Pathways
by
Guanyu Zhao, Xinying Yang, Yuehua Li, Muzi Li, Shiping Song, Jinghan Wang, Cong Liu, Zequn Dong, Zhenhua Gong and Junhui Liu
Viruses 2026, 18(9), 985; https://doi.org/10.3390/v18090985 - 8 Sep 2026
Abstract
Canine parvovirus 2 (CPV-2) is a major pathogen causing acute haemorrhagic enteritis and myocarditis in dogs. Since the discovery in the late 1970s, the original CPV-2 strain has evolved through VP2 mutations into new subtypes CPV-2a, 2b, and 2c. The viruses pose a
[...] Read more.
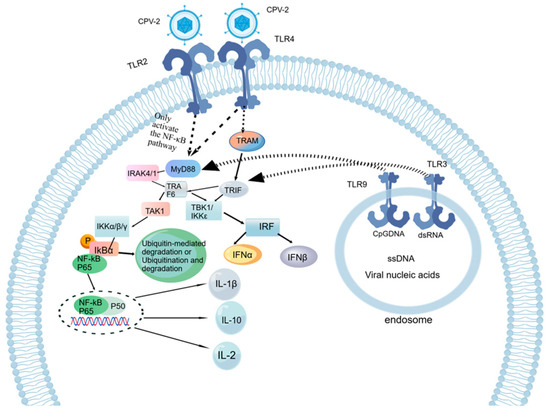
Canine parvovirus 2 (CPV-2) is a major pathogen causing acute haemorrhagic enteritis and myocarditis in dogs. Since the discovery in the late 1970s, the original CPV-2 strain has evolved through VP2 mutations into new subtypes CPV-2a, 2b, and 2c. The viruses pose a persistent threat to canine health worldwide. This review summarizes the innate immunity evasion tactics employed by CPV-2, focusing on the interactions of various viral proteins and host signaling pathways. CPV-2 regulates the activation and inhibition of Toll-like receptors (TLRs), thereby influencing the downstream MyD88/TRIF signaling pathway and modulating the activation of NF-κB and interferons. CPV-2 exerts bidirectional regulation of apoptosis, thereby modulating host innate immune responses and promoting viral replication. CPV-2 employs multiple strategies to evade the humoral and adaptive immune systems, including evolutionary escape through rapid replication and the accumulation of mutations. Clarifying the key mechanisms underlying the defective TLR activation, the immunomodulatory functions of CPV proteins, and cross-species transmission of CPV-2 among animals and the potential zoonotic risk to humans can provide a crucial theoretical basis for precise virus control, understanding immune evasion, and assessing zoonotic risks, with significant value for both vaccine development and basic research.
Full article
(This article belongs to the Special Issue Spotlight on Bocavirus and Other Parvoviruses, and Overlooked Respiratory Viruses)
►▼
Show Figures

Figure 1
Open AccessArticle
First European Identification of a Partial Tacheng Tick Virus 8-like Underscores the Hidden Burden of Tick-Borne Flavivirus-like Viruses
by
Guilherme Moreira, Eliane Silva, Joana Mourão and João R. Mesquita
Viruses 2026, 18(9), 984; https://doi.org/10.3390/v18090984 - 8 Sep 2026
Abstract
Tick-associated viruses are an underexplored component of global viromes, and highly divergent RNA viruses often remain undetected due to low abundance and sequence divergence. Tacheng tick virus 8 (TcTV8) is a poorly characterized RNA flavivirus-like virus originally reported from Dermacentor-associated ticks in China,
[...] Read more.
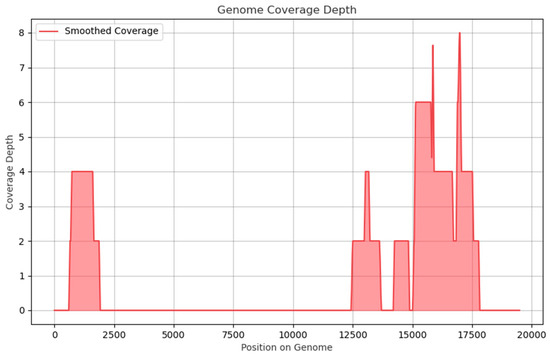
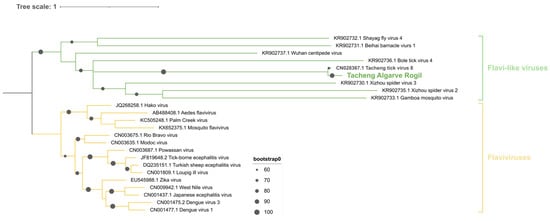
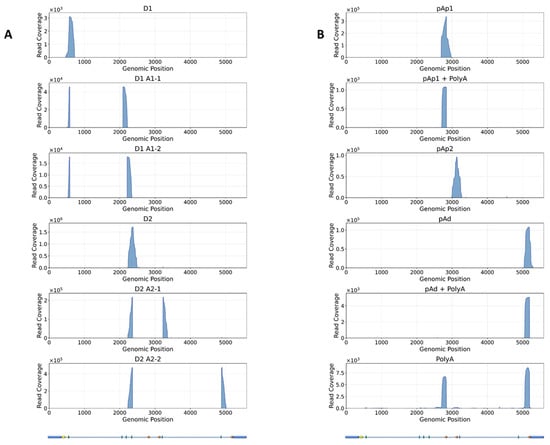
Tick-associated viruses are an underexplored component of global viromes, and highly divergent RNA viruses often remain undetected due to low abundance and sequence divergence. Tacheng tick virus 8 (TcTV8) is a poorly characterized RNA flavivirus-like virus originally reported from Dermacentor-associated ticks in China, with few subsequent reports and no confirmed detections outside Asia. Here, we report the detection and partial genomic characterization of a TcTV8-like virus in ticks collected from Portugal. Sequence-independent (SISPA) nanopore sequencing of individual ticks recovered three partial fragments (~2.7 kb, ~14% of the reference genome; ~30% of the reference covered at ≥1×), which shared high amino-acid identity with the TcTV8 polyprotein, including a methyltransferase-region domain. Phylogenetic analysis placed the Portuguese sequence within the TcTV8 lineage. Read-level classification and coverage analysis further support the presence of this virus in the positive sample. These findings represent, to our knowledge, the first detection of a TcTV8-like virus in European ticks, extending its known geographic range beyond Asia. The detection of this cryptic viral lineage highlights the need for broader tick virome surveillance to better understand the diversity, evolution, and ecology of flavivirus-like viruses.
Full article
(This article belongs to the Special Issue Synanthropy and Zoonotic Viral Spillover)
►▼
Show Figures

Figure 1
Open AccessArticle
Ebola Virus RNA Persists in Reproductive Tissues and Sperm of Male Ifnar1−/− Mice After Resolution of Viremia
by
Andrew L. Webb, Brayden G. Schindell, Kathy Frost, Geoff Soule, Stephanie A. Booth, David Safronetz and Jason Kindrachuk
Viruses 2026, 18(9), 983; https://doi.org/10.3390/v18090983 - 7 Sep 2026
Abstract
Ebola virus (EBOV) has been shown to persist in the reproductive tracts of male Ebola virus disease (EVD) survivors ranging from months to years following recovery, with shedding of EBOV in semen leading to chains of disease transmission. In the current study, we
[...] Read more.
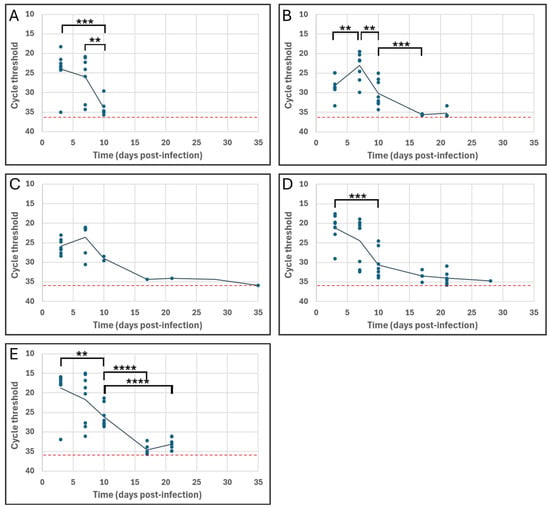
Ebola virus (EBOV) has been shown to persist in the reproductive tracts of male Ebola virus disease (EVD) survivors ranging from months to years following recovery, with shedding of EBOV in semen leading to chains of disease transmission. In the current study, we used a mouse model of EBOV infection to investigate the timing of virus infiltration of the male reproductive tissues, the extent of virus persistence in these tissues, and the impact of persistence on tissue health. We challenged male Ifnar−/− mice with wild-type EBOV and collected blood, liver, testis, epididymis, and sperm samples longitudinally for 35 days. All EBOV-challenged mice that were tested 3 days post-infection were viremic, and EBOV RNA was detected in reproductive samples up to 35 days post-infection. Epididymis and sperm samples were notable for detection of EBOV RNA at low cycle thresholds on day 3, and for high frequency of detection in samples up to 21 days post-infection. Testicular tissue histopathology was relatively mild during acute infection, but perturbations of interstitial tissue continued up to 35 days post-infection. Our findings support previous evidence of the risk of sexual transmission of persistent EBOV infections and highlight the potential impact of these infections on male fertility.
Full article
(This article belongs to the Special Issue Viruses in the Reproductive Tract)
►▼
Show Figures

Figure 1

Open AccessArticle
Real-World Effectiveness of Nirsevimab Against RSV-LRTI and Bronchiolitis Hospitalization: Results from the 2024–2025 Italian Universal Immunization Campaign in Newborns
by
Cinzia Auriti, Camilla Gizzi, Matteo Riccò, Roberto Bellù, Mario Giuffrè, Adele Fabiano, Caterina Cacace, Paola Cavicchioli, Armando Cuttano, Susanna Di Valerio, Cristiana Germini, Marcello Napolitano, Giulia Paviotti, Chiara Peila, Serafina Perrone, Simona Pesce, Gianfranco Scarpelli, Grazia Scutti, Giuseppina Spanedda, Alex Staffler, Massimo Agosti and on behalf of the Italian Working Group on RSV Prophylaxis of Italian Society of Neonatologyadd
Show full author list
remove
Hide full author list
Viruses 2026, 18(9), 982; https://doi.org/10.3390/v18090982 - 7 Sep 2026
Abstract
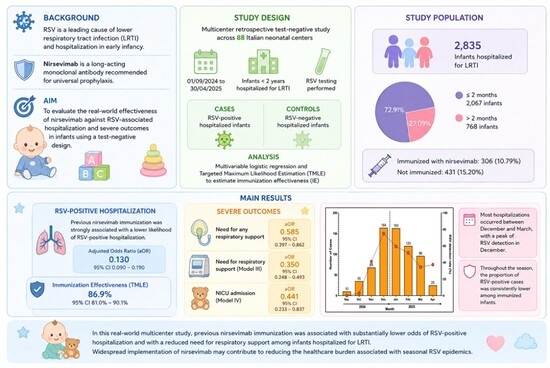
Respiratory syncytial virus (RSV) remains a leading cause of lower respiratory tract infection (LRTI) and hospitalization in early infancy. Following the introduction of universal nirsevimab prophylaxis in Italy during the 2024–2025 RSV season, we evaluated its real-world effectiveness against RSV-associated hospitalization and severe
[...] Read more.
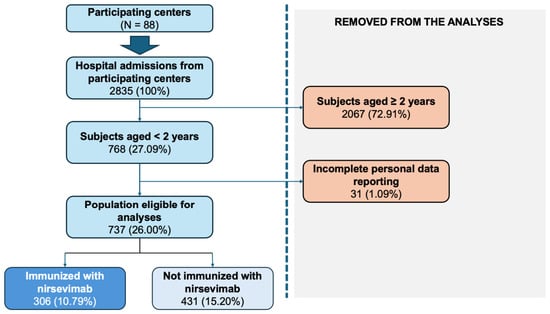
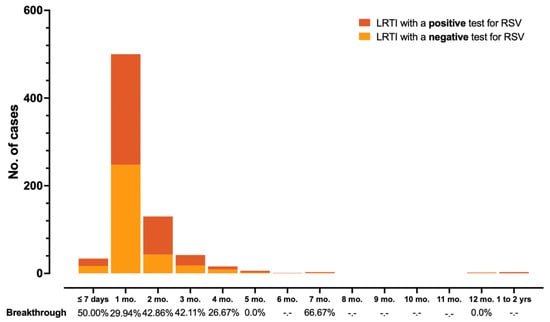
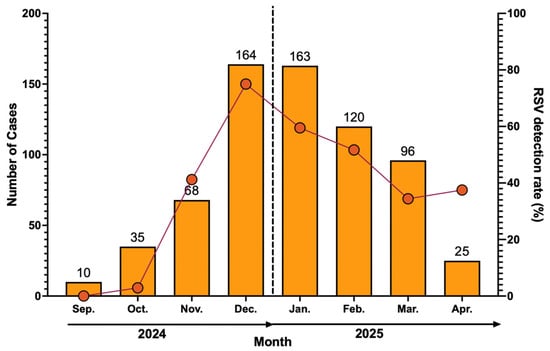
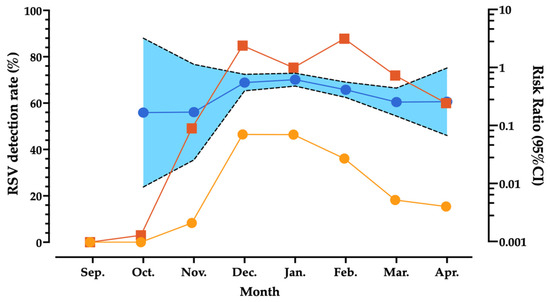
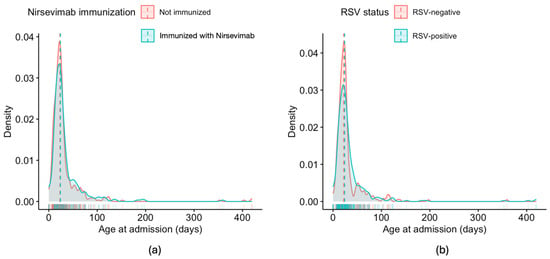
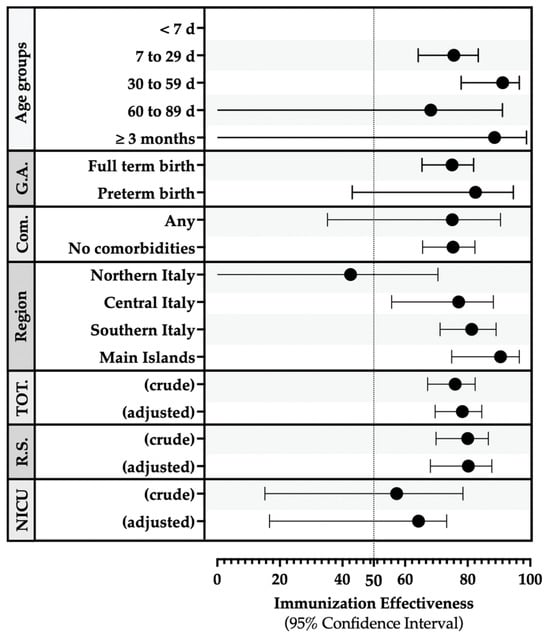
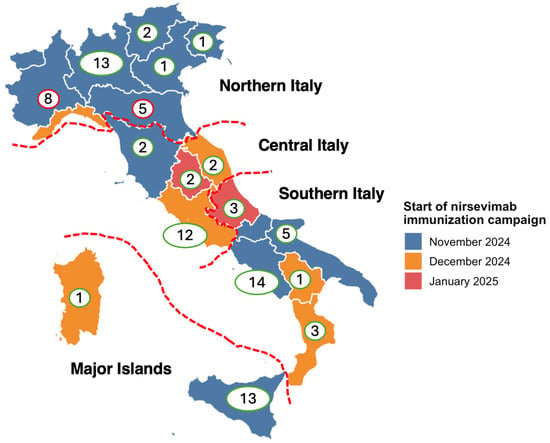
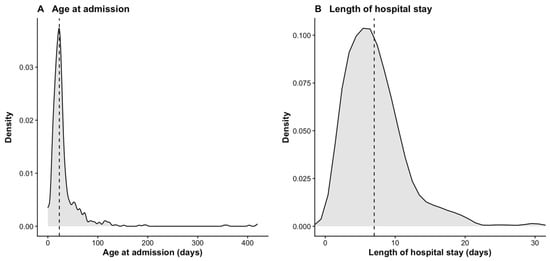
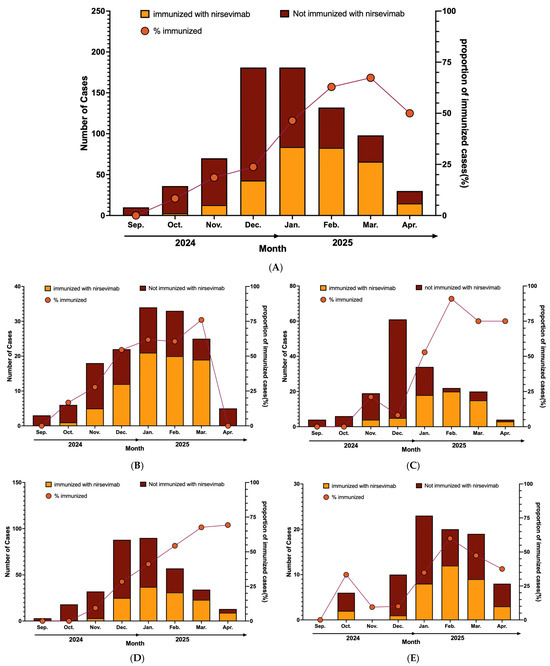
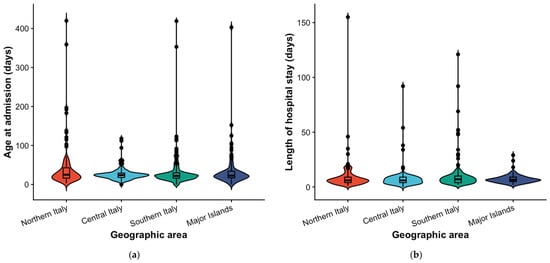
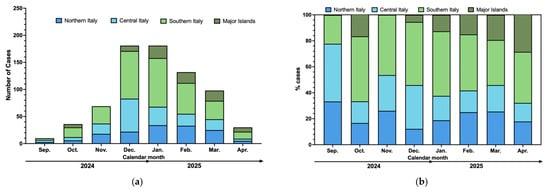
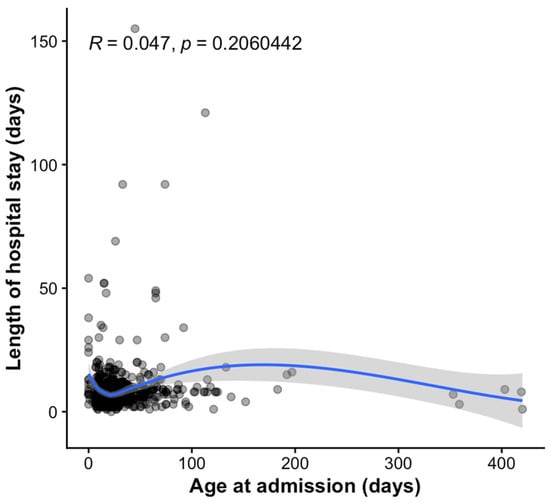
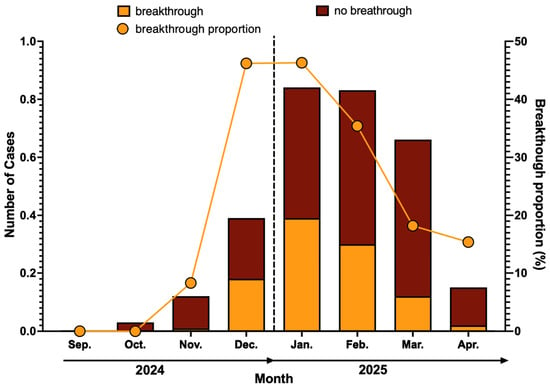
Respiratory syncytial virus (RSV) remains a leading cause of lower respiratory tract infection (LRTI) and hospitalization in early infancy. Following the introduction of universal nirsevimab prophylaxis in Italy during the 2024–2025 RSV season, we evaluated its real-world effectiveness against RSV-associated hospitalization and severe clinical outcomes within a test-negative design framework. We conducted a multicenter retrospective test-negative study across 88 Italian neonatal centers, including infants hospitalized for LRTI from 1 September 2024 to 30 April 2025. Clinical data were collected from hospital records using a standardized electronic questionnaire. RSV-positive hospitalized infants were considered cases, and RSV-negative hospitalized infants served as controls. Secondary outcomes included respiratory support and Neonatal Intensive Care Unit (NICU) admission. Effectiveness was estimated using multivariable logistic regression and targeted maximum likelihood estimation. Overall, 737 infants were included, of whom 306 (41.52%) had previously received nirsevimab. RSV infection was identified in 398 infants (54.00%), including 104/306 (33.99%) nirsevimab-immunized and 294/431 (68.21%) non-immunized infants (p < 0.001). Immunized infants also required respiratory support less frequently (55.23% vs. 70.07%; p < 0.001), with lower use of high-flow oxygen therapy (40.85% vs. 48.96%; p = 0.036) and Continuous Positive Airway Pressure (CPAP) (18.95% vs. 33.87%; p < 0.001). Within the test-negative design framework, adjusted effectiveness against RSV-associated hospitalization was estimated at 78.3%. Among hospitalized infants, previous nirsevimab immunization was additionally associated with adjusted relative reductions in the odds of respiratory support and NICU admission of 80.2% and 64.3%, respectively. In conclusion, among infants hospitalized for LRTI in routine Italian clinical practice, previous nirsevimab immunization was associated with lower odds of RSV positivity and a reduced need for respiratory support. Although these findings provide further evidence of the real-world effectiveness of nirsevimab against severe RSV disease, further studies are needed to provide direct estimates of hospitalization incidence in immunized and non-immunized source populations.
Full article
(This article belongs to the Section Human Virology and Viral Diseases)
►▼
Show Figures

Graphical abstract
Open AccessArticle
Elicitation of Potent H5N1- and H7N9-Neutralizing Antibody Responses in Rhesus Macaques by Sequential Heterologous Vaccination with mRNA and Adenoviral Vectors Encoding Full-Length Hemagglutinins
by
Brandon C. Rosen, Jack T. Mauter, Thomas B. Voigt, Giovana de Figueiredo Godoy, Emma L. Walker, Johan J. Louw, Noor Ghosh, Christakis Panayiotou, Aaron Yrizarry-Medina, Joshua Terao, Eva G. Rakasz, Matthew R. Reynolds, Dawn M. Dudley, Douglas S. Reed, David I. Watkins and Michael J. Ricciardi
Viruses 2026, 18(9), 981; https://doi.org/10.3390/v18090981 - 7 Sep 2026
Abstract
Highly pathogenic avian influenza (HPAI) represents an enormous pandemic risk amid the ongoing H5N1 panzootic. HPAI, defined by its high lethality in domestic poultry, includes the influenza A virus (IAV) H5N1 and H7N9 subtypes and has a 40–50% case fatality rate in humans.
[...] Read more.
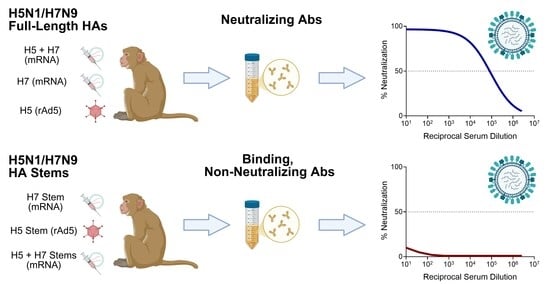
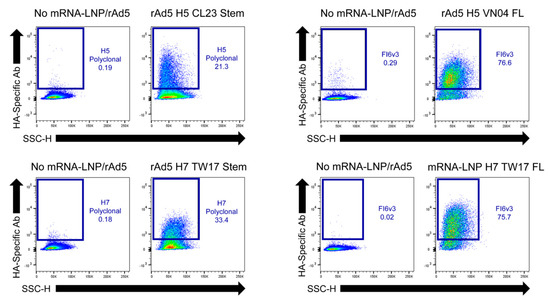
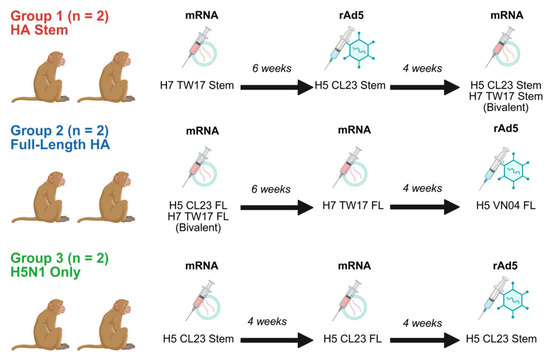
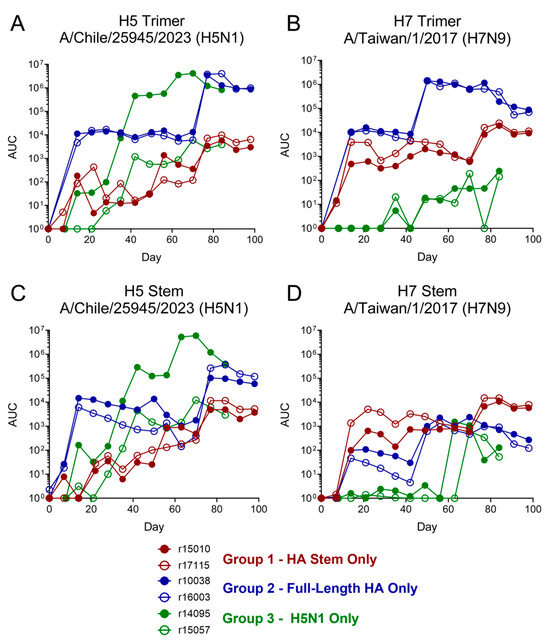
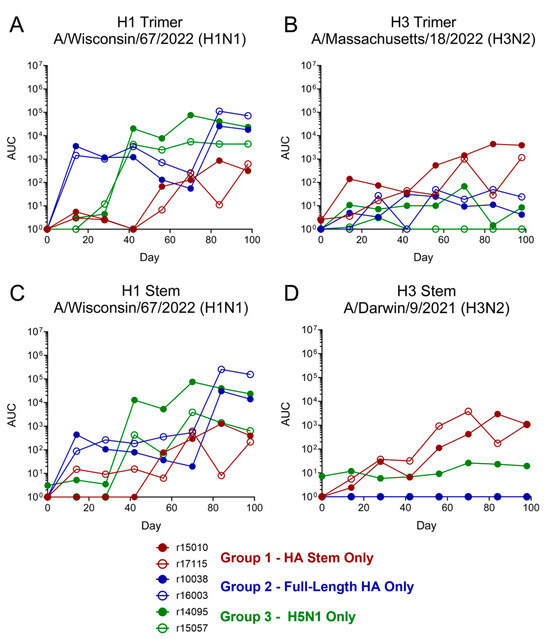
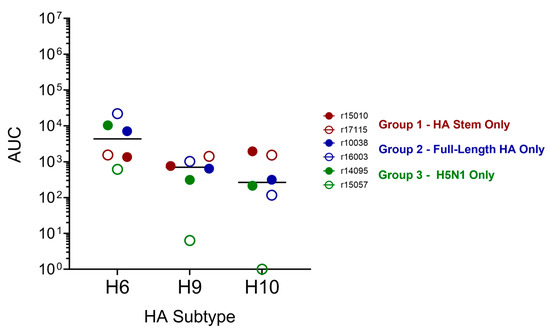
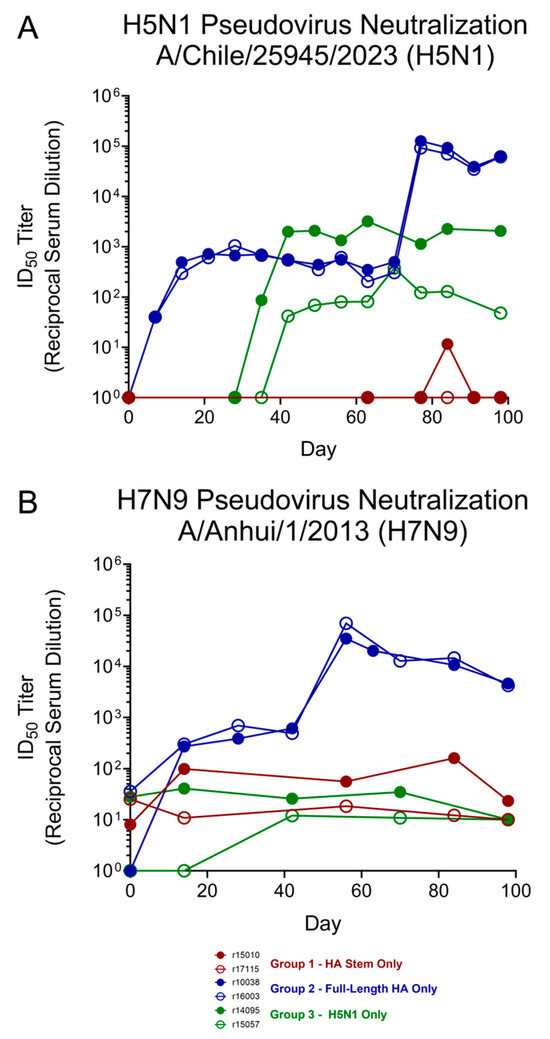
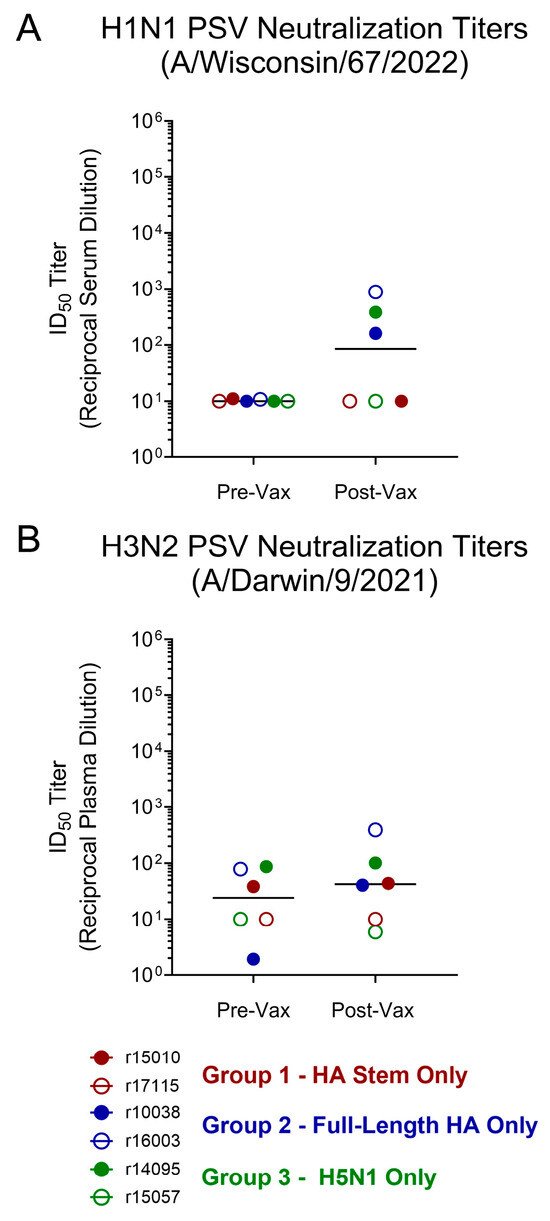
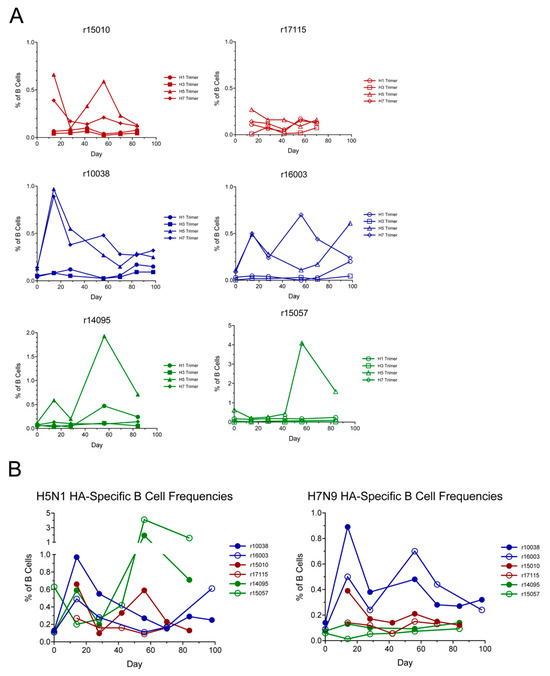
Highly pathogenic avian influenza (HPAI) represents an enormous pandemic risk amid the ongoing H5N1 panzootic. HPAI, defined by its high lethality in domestic poultry, includes the influenza A virus (IAV) H5N1 and H7N9 subtypes and has a 40–50% case fatality rate in humans. To facilitate the development of efficacious HPAI vaccination regimens and isolation of HPAI-neutralizing monoclonal antibodies (nmAbs) for prophylactic and therapeutic use, we performed a proof-of-concept pilot study wherein we developed an array of mRNA and adenovirus-vectored vaccines encoding HPAI hemagglutinins (HAs) and then assessed their immunogenicity in IAV-naïve Indian rhesus macaques (RMs). All RMs developed binding IgG recognizing both HPAI HAs. HPAI-nAbs were detected in RMs vaccinated with full-length HAs, but not in RMs vaccinated with HA stems alone. Potent HPAI neutralization activity was observed in two animals with serum ID50 titers of ~1:100,000 against H5N1 and ~1:50,000 against H7N9. Most RMs developed cross-reactive IgG recognizing the HAs of additional IAV subtypes, and HA-specific B cells were readily identifiable in vaccinee PBMCs by flow cytometric analysis. The results of our pilot study suggest that elicitation of potent HPAI-nAb responses is enhanced by vaccination with the HA head domain and that vaccination with the HA stem alone tends to elicit binding non-nAbs. Furthermore, use of our fluorophore-conjugated HA probes to identify HA-specific B cells could enable HPAI-nmAb isolation. Collectively, our findings could facilitate the development of novel vaccines and nmAb therapeutics leveraging the superior neutralization potency of HA head-specific Abs to prevent and treat HPAI infections in humans.
Full article
(This article belongs to the Section Viral Immunology, Vaccines, and Antivirals)
►▼
Show Figures

Graphical abstract
Open AccessReview
Emerging Biomarkers and Biotherapies for EBV-Associated Cancers
by
Maria C. White and Blossom Damania
Viruses 2026, 18(9), 980; https://doi.org/10.3390/v18090980 - 6 Sep 2026
Abstract
Epstein–Barr virus (EBV) is an oncogenic human gammaherpesvirus that is prevalent worldwide. Although over 95% of the world’s population harbors this pathogen, the vast majority of infections are asymptomatic, largely due to immune system involvement. However, in certain situations, such as when individuals
[...] Read more.
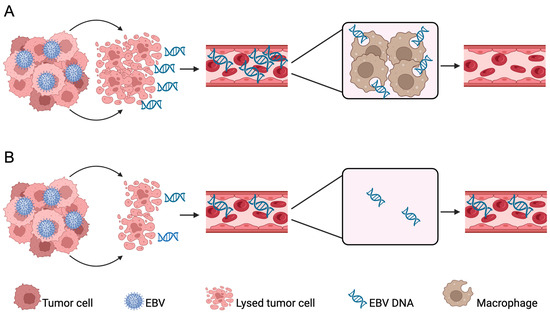
Epstein–Barr virus (EBV) is an oncogenic human gammaherpesvirus that is prevalent worldwide. Although over 95% of the world’s population harbors this pathogen, the vast majority of infections are asymptomatic, largely due to immune system involvement. However, in certain situations, such as when individuals are immunocompromised, the immune system can lose control of EBV suppression, leading to oncogenesis. While standard chemotherapeutic regimens can be effective against some EBV-associated cancers, these treatments can be toxic to some patients, and disease relapse and/or drug resistance can develop. Additionally, no human vaccine currently exists for EBV. In recent years, biotherapy has become a promising avenue for the clinical management of EBV-positive cancers. As early detection is often correlated with a more positive prognosis, identification of reliable EBV biomarkers has also come into focus. In this review, we summarize advancements made over the last five years in immunotherapy use for EBV-positive cancers and in EBV biomarker identification to detect both EBV infection and biotherapy response.
Full article
(This article belongs to the Special Issue Immune Evasion and Viral Carcinogenesis)
►▼
Show Figures

Figure 1
Open AccessArticle
Powassan Virus Infects Mouse Testes and Reveals a Protective Role for Phagocytic Cells
by
E. Eldridge Hager-Soto, Esteban Arroyave, Eduardo Eyzaguirre, Omar A. Saldarriaga, Alexander N. Freiberg and Shannan L. Rossi
Viruses 2026, 18(9), 979; https://doi.org/10.3390/v18090979 - 5 Sep 2026
Abstract
Powassan virus (POWV) is an emerging tick-borne orthoflavivirus with consistently increasing annual prevalence. Though understood as a vector-transmitted virus associated with neurological disease, a 2016 fatal case of an immunocompromised 63-year-old man reported testicular pain with postmortem detection of viral antigen in the
[...] Read more.
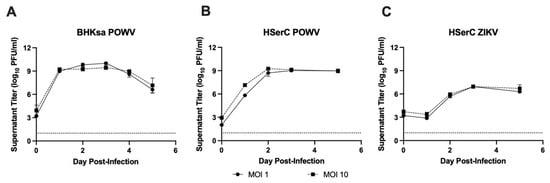
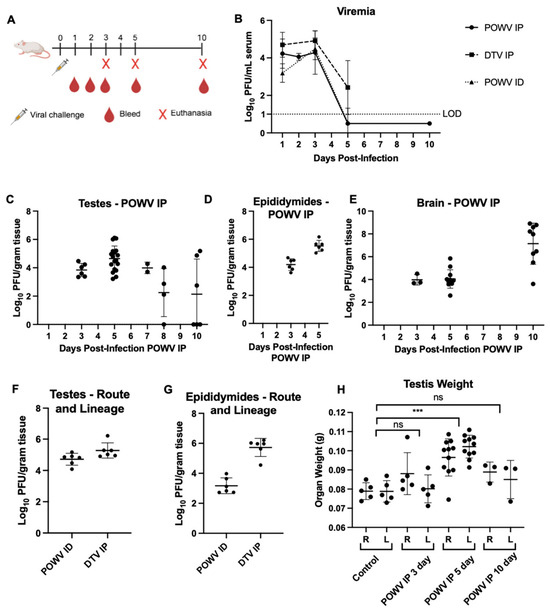
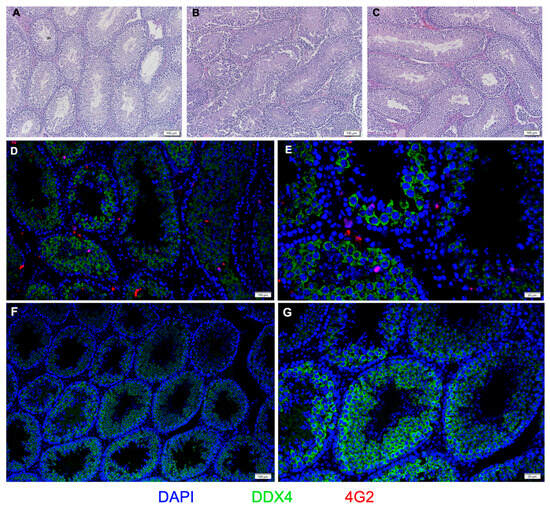
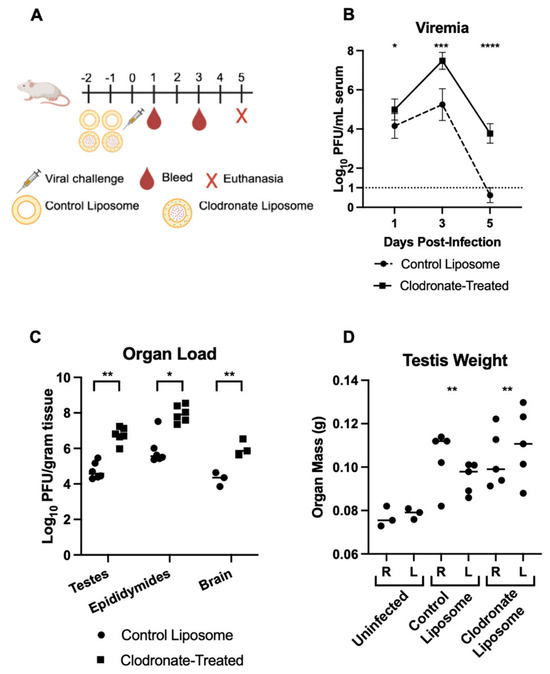
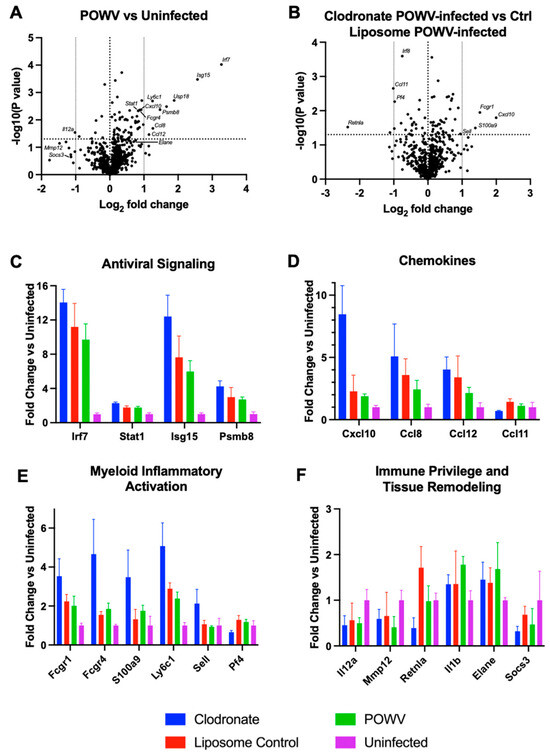
Powassan virus (POWV) is an emerging tick-borne orthoflavivirus with consistently increasing annual prevalence. Though understood as a vector-transmitted virus associated with neurological disease, a 2016 fatal case of an immunocompromised 63-year-old man reported testicular pain with postmortem detection of viral antigen in the testes, suggesting an altered tropism. The extent to which POWV can infect testes remains unclear. We used a BALB/c mouse model of POWV to elucidate the potential of POWV to establish infection in testes and model a potential mechanism of testicular tropism. Here, we report that POWV productively infects the male reproductive tract of mice. We find that POWV consistently infects mouse testes and epididymides at 3 and 5 days post-infection (dpi) with peak titers measuring 6.1 log10 PFU/gram of tissue in testes. Furthermore, we find that at 5 dpi, POWV-infected mice testes are significantly higher in mass than uninfected controls. Due to macrophages being implicated in driving testicular pathogenesis in Zika virus, a related orthoflavivirus, we depleted phagocytic cells using clodronate liposomes prior to POWV infection and compared them to control liposome-treated mice. Depletion of phagocytic cells was associated with higher and sustained viremia and significantly higher viral organ burden in the male reproductive tract of mice. Gene expression analysis revealed a robust antiviral, inflammatory, and chemokine response in the testes that was further amplified following phagocytic-cell depletion. These findings implicate the male reproductive tract as a previously underrecognized target of POWV disease and identify phagocytic cells as key in limiting viral infection of testes in mice.
Full article
(This article belongs to the Special Issue Pathogenesis and Persistence in Flavivirus Infections: Mechanisms, Biomarkers, and Clinical Implications)
►▼
Show Figures

Figure 1
Open AccessArticle
Epidemiological Characteristics and Evolutionary Characterization of Human Metapneumovirus in Jiaxing, China
by
Yamei Zhou, Yanqian Wu, Peiyan He, Yong Yan, Ganglin Ren, Xiaofei Zhang, Yin Song and Guoyin Zhu
Viruses 2026, 18(9), 978; https://doi.org/10.3390/v18090978 - 4 Sep 2026
Abstract
Human metapneumovirus (hMPV) represents a leading cause of both upper and lower respiratory tract infections among children and adults globally. To investigate the prevalence and evolution of hMPV in the Jiaxing area of China between 2023 and 2025, we screened 3600 pharyngeal swab
[...] Read more.
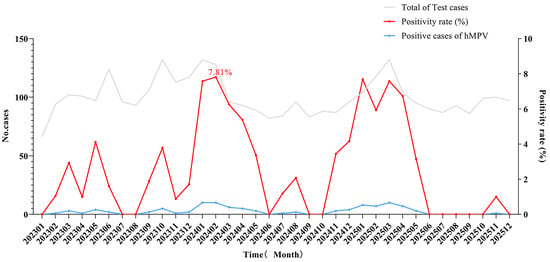
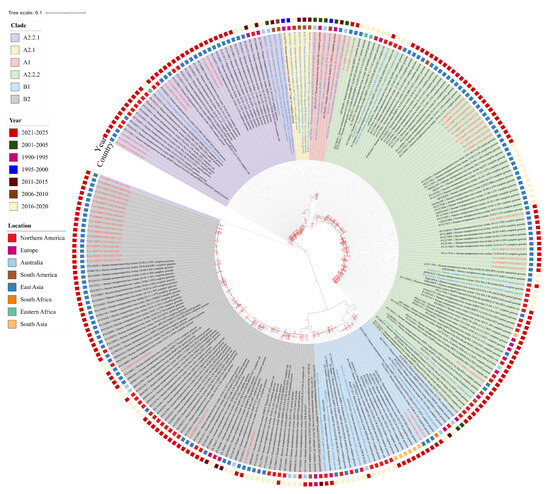
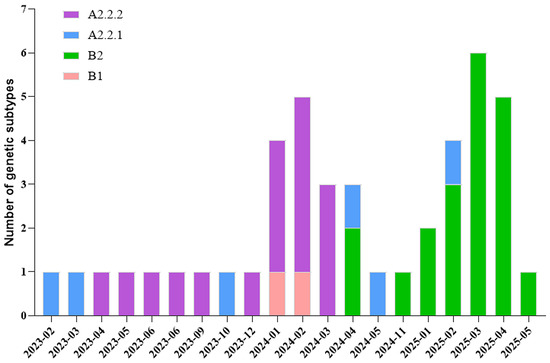
Human metapneumovirus (hMPV) represents a leading cause of both upper and lower respiratory tract infections among children and adults globally. To investigate the prevalence and evolution of hMPV in the Jiaxing area of China between 2023 and 2025, we screened 3600 pharyngeal swab specimens by real-time PCR, identified 101 positives, and obtained genomic sequences of 48 viral isolates using high-throughput sequencing. Using the sequencing data, we reconstructed a phylogenetic tree and examined amino acid substitutions. The epidemiological analysis revealed an overall hMPV positivity rate of 2.81% (101/3600) in Jiaxing during 2023–2025. Although positive cases were detected across all age groups, they were mainly children, with no significant difference between genders. Regarding seasonal patterns, the peak of hMPV activity occurred predominantly during winter and spring. Over the study period, four genotypes co-circulated, in the order of B2 (41.67%), A2.2.2 (37.50%), A2.2.1 (16.67%), and B1 (4.17%). Further phylogenetic analysis showed that the B1 strains from Jiaxing clustered primarily with those from Beijing, China, while B2, A2.2.1, and A2.2.2 strains were more closely related to strains from the United States and Beijing. Of note, an A2c111nt-dup variant was identified in Jiaxing in 2023. Starting from November 2024, the prevailing genotype transitioned from A2.2.1/A2.2.2 to B2, and B2 emerged as the absolutely dominant strain by 2025. In comparison with earlier circulating strains, several amino acid substitutions have accumulated in current isolates, such as T223N, D280N, I392T, R396Q, S444N, K450R, and T521A in the F protein of B2 strains. Furthermore, the G, L, P, and SH proteins also displayed temporally patterned amino acid replacements. While the biological significance of these mutations is yet to be determined, these results highlight the public health importance of ongoing hMPV surveillance and dynamic monitoring of its genetic evolution.
Full article
(This article belongs to the Section Human Virology and Viral Diseases)
►▼
Show Figures

Figure 1
Open AccessArticle
Global Patterns of Geographic Distribution, Temporal Trends, Host Spectrum, and Molecular Variation of H9N2 Avian Influenza Virus, 1966–2023
by
Lei Chang, Xiangyan Ren, Lebin Han, Chenlu Xia and Yuzhong Zhao
Viruses 2026, 18(9), 977; https://doi.org/10.3390/v18090977 - 4 Sep 2026
Abstract
The H9N2 avian influenza virus (AIV) is currently widespread globally and poses a serious threat to the poultry industry and public health; however, its global epidemiological characteristics and key molecular mutation patterns have not yet been systematically elucidated. Based on the NCBI Influenza
[...] Read more.
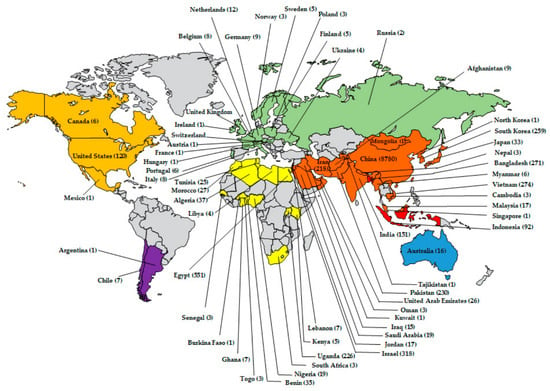
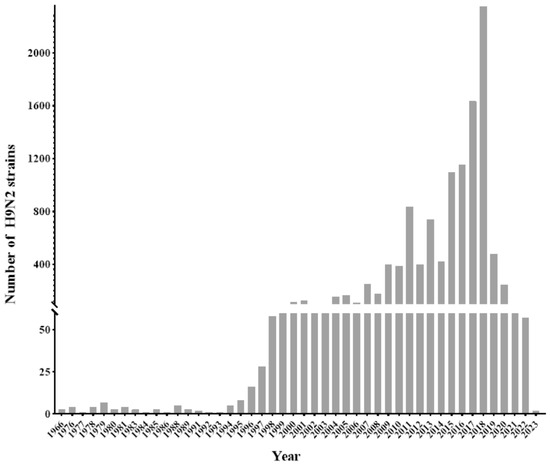
The H9N2 avian influenza virus (AIV) is currently widespread globally and poses a serious threat to the poultry industry and public health; however, its global epidemiological characteristics and key molecular mutation patterns have not yet been systematically elucidated. Based on the NCBI Influenza Virus Resource Database, this study collected relevant data on the H9N2 AIV from around the world between 1966 and 2023. It conducted a comprehensive analysis of the geographic distribution of 11,933 strains, the temporal distribution of 11,824 strains, the host sources of 11,756 strains, and the genetic polymorphisms at key functional sites. The results showed that 90.14% of H9N2 AIV sequences in the dataset originated from Asia, with China contributing the largest number of sequences. The global sequence availability pattern underwent four phases: sporadic sequence availability, gradual increase, high sequence availability, and subsequent decline, with 2007–2018 representing a period of relatively high availability of publicly accessible sequences. Host analysis indicated that H9N2 AIV sequences were predominantly derived from poultry-related hosts, with chickens being the main host source in the dataset, and available sequences have also been reported from various mammals, including pigs and humans, as well as environmental media. Molecular analysis revealed that receptor-binding-related sites in the HA protein, such as Q226L and I155T, have become dominant variants, while mammalian-adaptive mutations such as PB2 E627K and D701N remain at low frequencies; M2 S31N is widely prevalent, whereas the detection rate of NA drug-resistance-associated mutations is low. This study systematically reveals the global distribution patterns of available H9N2 AIV sequences, host distribution characteristics, and key molecular mutation patterns of the H9N2 AIV, providing a scientific basis for cross-host transmission risk assessment, molecular surveillance, and targeted prevention and control.
Full article
(This article belongs to the Special Issue Molecular Epidemiology, Evolution, and Transmission of Avian Influenza Viruses: 2nd Edition)
►▼
Show Figures

Figure 1
Open AccessArticle
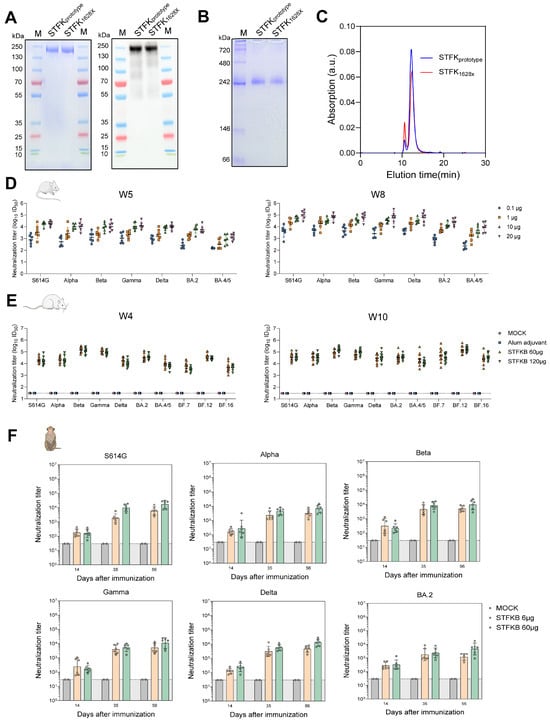
An Engineered Spike Immunogen Drives Broad Protection of a Bivalent Recombinant Protein COVID-19 Vaccine in Animal Models
by
Zikang Wang, Jiahua Gao, Ruojing Bai, Lunzhi Yuan, Huilin Guo, Shaojuan Wang, Youfeng Wang, Jiayi Wu, Qingfang Bu, Huiyu Guo, Yunda Hong, Jian Ma, Kai Wang, Wenjie Guo, Yanyan Liu, Zhitao Weng, Tianying Zhang, Ningshao Xia, Quan Yuan, Yali Zhang and Yangtao Wuadd
Show full author list
remove
Hide full author list
Viruses 2026, 18(9), 976; https://doi.org/10.3390/v18090976 - 4 Sep 2026
Abstract
The continued evolution of SARS-CoV-2 has reduced the protective effectiveness of first-generation vaccines and underscores the need for broadly reactive next-generation vaccine candidates. Here, we evaluated STFKB, an alum-adjuvanted bivalent recombinant protein vaccine composed of the monomeric Spike (STFKprototype) and the
[...] Read more.
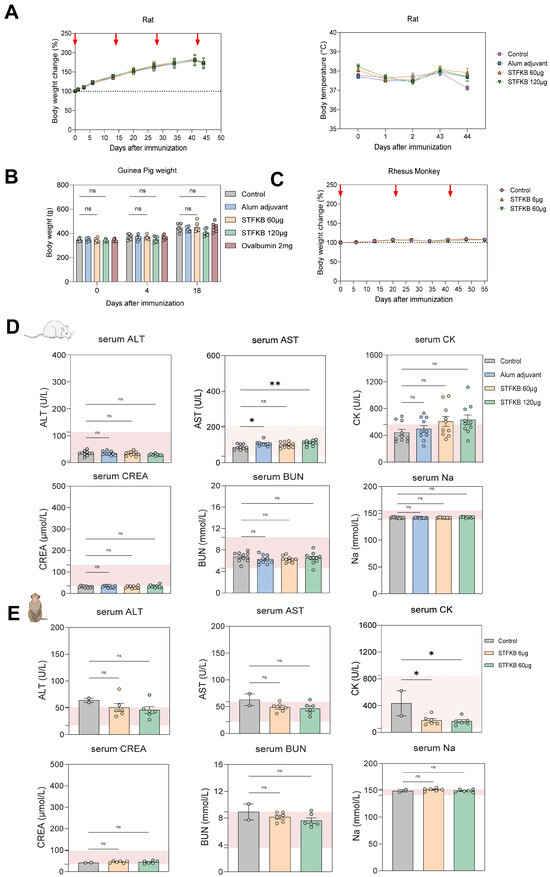
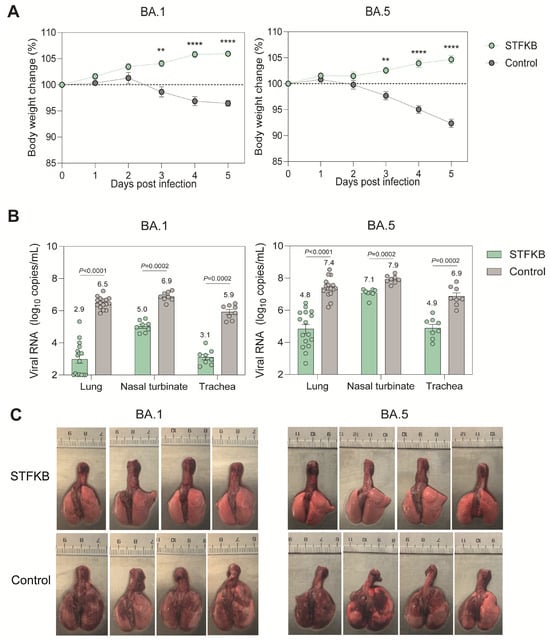
The continued evolution of SARS-CoV-2 has reduced the protective effectiveness of first-generation vaccines and underscores the need for broadly reactive next-generation vaccine candidates. Here, we evaluated STFKB, an alum-adjuvanted bivalent recombinant protein vaccine composed of the monomeric Spike (STFKprototype) and the engineered Spike (STFK1628X). We assessed the immunogenicity, tolerability, and protective efficacy of STFKB in mice, rats, guinea pigs, rhesus macaques, and Syrian hamsters. STFKB induced robust STFK- and STFK1628X-specific antibody responses and broadly reactive neutralizing antibodies against multiple SARS-CoV-2 variants in the tested animal models. In hamster challenge studies, STFKB vaccination protected animals from Omicron BA.1 and BA.5 challenge, as shown by reduced body-weight loss, lower viral RNA loads in respiratory tissues, and improved gross lung pathology. Across the tested preclinical models, STFKB was well tolerated, with no vaccine-related overt toxicity observed under the study conditions. These findings support the rationale and translational potential of the bivalent vaccine strategy proposed in this study.
Full article
(This article belongs to the Section Coronaviruses)
►▼
Show Figures

Figure 1
Journal Menu
► ▼ Journal Menu-
- Viruses Home
- Aims & Scope
- Editorial Board
- Reviewer Board
- Topical Advisory Panel
- Instructions for Authors
- Special Issues
- Topics
- Sections & Collections
- Article Processing Charge
- Indexing & Archiving
- Editor’s Choice Articles
- Most Cited & Viewed
- Journal Statistics
- Journal History
- Journal Awards
- Society Collaborations
- Conferences
- Editorial Office
Journal Browser
► ▼ Journal BrowserHighly Accessed Articles
Latest Books
E-Mail Alert
News
Topics
Topic in
Animals, Encyclopedia, Pathogens, TropicalMed, Veterinary Sciences, Viruses, Zoonotic Diseases
Emerging and Resurging Viruses: Zoonotic Threats in a Changing World
Topic Editors: Levon Abrahamyan, Francesco VairoDeadline: 31 March 2027
Topic in
Biomedicines, Cancers, IJMS, Vaccines, Viruses, JMP
Advances in HPV-Driven Head and Neck Cancer: Pathogenesis, Treatment, and Prevention
Topic Editors: Shilpi Gupta, Bhudev Chandra Das, Prabhat KumarDeadline: 20 April 2027
Topic in
AppliedMath, BioMedInformatics, Infectious Disease Reports, Mathematics, Viruses
Modeling, Dynamics, and Control of Infectious Diseases via Complex Network Approaches
Topic Editors: Wei Yao, Hongying ShuDeadline: 30 September 2027
Topic in
Antibiotics, Diseases, Vaccines, Viruses, Epidemiologia
HIV Prevention and Treatment: A Guide to Health and Safety
Topic Editors: Olanrewaju Oladimeji, Martha ChadyiwaDeadline: 31 December 2027
Conferences
Special Issues
Special Issue in
Viruses
Hepatitis B Virus—Mechanisms of Persistence and Progress Towards a Cure
Guest Editors: Qiong Zhao, Yongzhen LiuDeadline: 16 September 2026
Special Issue in
Viruses
The Structure and Function of Flavivirus Genes and Proteins
Guest Editors: Tadahisa Teramoto, Kay ChoiDeadline: 30 September 2026
Special Issue in
Viruses
Viruses in the Reproductive Tract
Guest Editors: Mimi Ghosh, Raina Nakova Fichorova, Christina FarrDeadline: 30 September 2026
Special Issue in
Viruses
Imaging Neurotropic Viruses
Guest Editors: Elena Criscuolo, Salla MattolaDeadline: 30 September 2026
Topical Collections
Topical Collection in
Viruses
Poxviruses
Collection Editors: Giliane de Souza Trindade, Galileu Barbosa Costa, Flavio Guimaraes da Fonseca
Topical Collection in
Viruses
Phage Therapy
Collection Editors: Nina Chanishvili, Jean-Paul Pirnay, Mikael Skurnik