-
Beyond the Tropics: Clinical Presentation and Epidemiology of Travel-Associated Dengue Fever in a Non-Endemic European Setting -
Methane Production on Mars-Relevant Clay Minerals -
Identification and Genomic Localization of the cpe Gene in Clostridium perfringens -
A Structural View of Influenza Virus Ribonucleoprotein Complex and Its Functions
Journal Description
Microorganisms
Microorganisms
is a scientific, peer-reviewed, open access journal of microbiology, published monthly online by MDPI. The Hellenic Society Mikrobiokosmos (MBK), the Spanish Society for Nitrogen Fixation (SEFIN) and the Society for Microbial Ecology and Disease (SOMED) are affiliated with Microorganisms, and their members receive a discount on the article processing charges.
- Open Access— free for readers, with article processing charges (APC) paid by authors or their institutions.
- High Visibility: indexed within Scopus, SCIE (Web of Science), PubMed, PMC, PubAg, CAPlus / SciFinder, AGRIS, and other databases.
- Journal Rank: JCR - Q2 (Microbiology) / CiteScore - Q1 (Virology)
- Rapid Publication: manuscripts are peer-reviewed and a first decision is provided to authors approximately 16.5 days after submission; acceptance to publication is undertaken in 3.3 days (median values for papers published in this journal in the first half of 2026).
- Recognition of Reviewers: Reviewers whose reports are timely and of high quality receive an APC discount voucher for a future publication in an MDPI journal. Become a reviewer.
- Companion journal for Microorganisms include: Applied Microbiology and Bacteria.
- Journal Cluster of Microbiology: Acta Microbiologica Hellenica, Applied Microbiology, Bacteria, Journal of Fungi, Microorganisms, Microbiology Research, Pathogens, Viruses, Fermentation and Germs.
Impact Factor:
4.7 (2025);
5-Year Impact Factor:
5.1 (2025)
Latest Articles
Seasonal Variations in Microbial Community Structure and Function in the Waters Along the Yangtze-to-Huaihe Water Diversion Project According to Metagenomics
Microorganisms 2026, 14(9), 2016; https://doi.org/10.3390/microorganisms14092016 - 10 Sep 2026
Abstract
Water diversion projects can alleviate the uneven spatiotemporal distribution of water resources, but they may also impact functions of aquatic ecosystems in the waters along the route. Despite their importance, the temporal and spatial changes in multi-domain microbial community structure and function along
[...] Read more.
Water diversion projects can alleviate the uneven spatiotemporal distribution of water resources, but they may also impact functions of aquatic ecosystems in the waters along the route. Despite their importance, the temporal and spatial changes in multi-domain microbial community structure and function along the route remain poorly understood. Metagenomics was employed to investigate the community structure and function of bacteria, archaea, and fungi in the aquatic environments along the Yangtze-to-Huaihe water diversion project in winter and summer seasons. The results showed that seasonal variations may drive a trade-off in the species diversity of bacterial and fungal communities. Seasonal variations altered microbial communities (especially for the bacteria), and exerted a greater influence on community structure than spatial factors. Microbial community composition was more sensitive to seasonal fluctuations than functional genes. The species spatial turnover played a dominant role in shaping microbial communities (especially for winter) in both seasons. Archaea, bacteria, fungi and KEGG functional genes all exhibited a positive correlation with some environmental factors in summer but not in winter. PLS-SEM indicated that water quality and microbial composition directly significantly impacted functional genes. This study offers a theoretical basis for maintaining the stability of water ecological microorganisms in water transfer projects.
Full article
(This article belongs to the Section Environmental Microbiology)
Open AccessArticle
Metagenomic Insights into Microbial Functional Potential Associated with Soil Carbon, Nitrogen, and Phosphorus Cycling Along an Elevational Gradient in a Warm-Temperate Forest
by
Jingjing Wang, Siyuan Huangfu, Ruochen Li, Haibo Li, Hongyi He, Biaobing Chang, Huinan Ma, Haoqin Ma, Jiaxin Zhang, Ruohong Hou, Houjuan Song and Xiuqing Yang
Microorganisms 2026, 14(9), 2015; https://doi.org/10.3390/microorganisms14092015 - 10 Sep 2026
Abstract
Soil microbial functional potential is crucial to maintaining forest productivity and ecosystem functions. However, how microbially mediated soil nutrient cycling responds to environmental changes, particularly those caused by variations in elevation, remains poorly understood. Using the natural temperature gradient in a temperate mountain
[...] Read more.
Soil microbial functional potential is crucial to maintaining forest productivity and ecosystem functions. However, how microbially mediated soil nutrient cycling responds to environmental changes, particularly those caused by variations in elevation, remains poorly understood. Using the natural temperature gradient in a temperate mountain forest, this study investigated the differences in functional microbial groups and functional genes involved in soil carbon, nitrogen and phosphorus cycling along the elevation gradient, and analyzed the associations between environmental factors and these differences. The results showed that the low-elevation gradient (LE) had significantly higher abundances of genes involved in carbon degradation (pfkC, pgi1, and LSC1) but significantly lower abundances of those involved in carbon fixation (K18602, K18603, and K18604). Compared with the high-elevation gradient (HE), the LE had a significantly higher abundance of the nitrogen-cycle gene involved in organic degradation and synthesis (nao), but significantly lower abundances of denitrification (norB) and dissimilatory nitrate reduction genes (narG, narI, and napC). The abundances of the key genes involved in phosphorus metabolism (aphA and purO) were significantly higher at HE than at LE, whereas the abundance of the key gene associated with phosphorus transport (phnT) was significantly lower. The composition of the microbial community at the phylum level involved in carbon, nitrogen and phosphorus cycling at different elevations was similar, but the relative abundance of Thermoproteota and Nitrospirota increased significantly at HE. The annual average temperature, pH and carbon acquisition enzymes (β-glucosidase and β-D-cellobiosidase) were significantly associated with microbial community composition and functional genes related to carbon, nitrogen and phosphorus cycles. Additionally, genes involved in the carbon, nitrogen and phosphorus cycles were closely related through synergy and antagonism, especially the metabolic pathways encoded by purO, phnT and nrfA. These results provide metagenomic insights into the response patterns of microbial functional potential associated with soil carbon, nitrogen, and phosphorus cycling along an elevational gradient in a warm-temperate forest.
Full article
(This article belongs to the Section Environmental Microbiology)
►▼
Show Figures

Figure 1
Open AccessFeature PaperArticle
Growth-Phase-Dependent Shift in GABA Biosynthetic Pathways Under Temperature Stress in Isochrysis zhanjiangensis
by
Jiansen Luo, Lin Zhang, Jichang Han, Yumeng Wang, Jiaxin Yu, Jingbo Fan, Lulu Wang, Jiayi Cao, Kehou Pan and Jilin Xu
Microorganisms 2026, 14(9), 2014; https://doi.org/10.3390/microorganisms14092014 - 10 Sep 2026
Abstract
Temperature stress is a major constraint on the productivity of microalgae used in aquaculture. γ-Aminobutyric acid (GABA) is well-established as a key player in the stress tolerance of higher plants, yet its role in microalgae remains largely unexplored. Here, we examined the effects
[...] Read more.
Temperature stress is a major constraint on the productivity of microalgae used in aquaculture. γ-Aminobutyric acid (GABA) is well-established as a key player in the stress tolerance of higher plants, yet its role in microalgae remains largely unexplored. Here, we examined the effects of low (15 °C), optimal (25 °C), and high (35 °C) temperatures on the GABA shunt in Isochrysis zhanjiangensis during the initial and mid-exponential growth phases. The results demonstrated that temperature stress significantly inhibited cell growth and photosynthetic efficiency (assessed by Fv/Fm and Fv’/Fm’), with soluble protein decreasing and soluble sugar accumulating. During the initial exponential phase, both low and high temperature stress triggered marked GABA accumulation, accompanied by coordinated increases in glutamate decarboxylase (GAD) and diamine oxidase (DAO) activities. Interestingly, the transcript levels of IzGAD and IzDAO decreased under these conditions, suggesting that GABA accumulation at this stage is predominantly governed by post-translational activation rather than transcriptional upregulation. Upon entry into the mid-exponential phase, a distinct phase-dependent shift in GABA biosynthetic regulation emerged. Under low temperature stress, GAD activity and IzGAD expression were both suppressed, whereas DAO activity and IzDAO transcripts increased significantly, indicating the transition to DAO-mediated GABA production as the dominant route. Under high temperature stress, both GAD and DAO activities increased, yet their corresponding gene transcription remained repressed, revealing a persistent asynchrony between enzyme activities and gene expression across both phases. Meanwhile, the expression of catabolic genes (IzGABA-T, IzSSADH1, and IzSSADH2) was consistently downregulated, further facilitating the net accumulation of GABA. Promoter analysis revealed multiple stress- and hormone-responsive cis-elements in these genes, implying a complex regulatory network. Collectively, our findings uncover a growth-phase-dependent reconfiguration of GABA biosynthetic pathways in I. zhanjiangensis under temperature stress. These insights provide a mechanistic basis for strain-specific temperature management in aquaculture applications.
Full article
(This article belongs to the Special Issue Molecular Mechanisms of Algal and Microbial Adaptation to Extreme Temperature Conditions)
►▼
Show Figures

Figure 1
Open AccessReview
Innovative Applications of Artificial Intelligence in Bacteriophage Research: A New Chapter in Future Medicine
by
Dapeng Yang, Xin Yuan and Yubao Li
Microorganisms 2026, 14(9), 2013; https://doi.org/10.3390/microorganisms14092013 - 10 Sep 2026
Abstract
As the crisis of antibiotic resistance escalates, phage therapy has regained attention as an alternative strategy. Artificial intelligence (AI) technologies offer new avenues to overcome the bottlenecks inherent in traditional bacteriophage research. This review summarizes the multi-dimensional innovative applications of machine learning, deep
[...] Read more.
As the crisis of antibiotic resistance escalates, phage therapy has regained attention as an alternative strategy. Artificial intelligence (AI) technologies offer new avenues to overcome the bottlenecks inherent in traditional bacteriophage research. This review summarizes the multi-dimensional innovative applications of machine learning, deep learning, and large biological models in phage studies. In the fields of phage recognition and genomics, support vector machines (SVMs), convolutional neural networks (CNNs), and pre-trained protein language models can all achieve recognition accuracy rates of over 90%. Furthermore, tools such as DeepHost and VirSorter2 can efficiently identify phage sequences, annotate functional genes, and predict hosts at the species or strain levels. For clinical translation, AI integrates patient characteristics, bacterial phenotypes, and phage profiles to customize cocktail regimens for individualized phage therapy. Graph neural network-based models like DeepPBI-KG integrate multi-omics knowledge graphs to precisely predict phage-host interactions (PHIs), whereas agent-based simulation and defense protein predictors forecast phage resistance evolution. Additionally, generative AI can support the de novo design of functional phage genomes and mine massive unannotated virome dark matter. Nevertheless, this cross-disciplinary field faces significant constraints, including uneven and biased sequencing datasets, insufficient model interpretability, and dual-use biosafety ethical risks accompanied by unclear algorithm accountability and incomplete global supervision systems. Future research should optimize standardized multimodal databases, develop explainable AI algorithms, and establish cross-disciplinary ethical governance frameworks to facilitate closed-loop verification between computational prediction and wet-lab experiments. In conclusion, the deep integration of AI and phage biology provides revolutionary strategies to tackle multidrug-resistant infections and advances the clinical transformation of phage precision medicine.
Full article
(This article belongs to the Special Issue Artificial Intelligence as a Tool for Combating Antimicrobial Resistance)
►▼
Show Figures

Figure 1
Open AccessArticle
Seasonal Dynamics of ‘Candidatus Phytoplasma prunorum’ in Selected Prunus Species Revealed by Quantitative PCR
by
Peter Morvay, Tomáš Kiss, Ivo Ondrášek, Erika Slámová, Eliška Zezulová and Tomáš Nečas
Microorganisms 2026, 14(9), 2012; https://doi.org/10.3390/microorganisms14092012 - 10 Sep 2026
Abstract
The distribution of ‘Candidatus Phytoplasma prunorum’, a phloem-limited pathogen, varies among plant tissues during the year, but knowledge about quantitative comparisons of its presence in roots and above-ground tissues across Prunus species remains limited. The variation in ‘Ca. P. prunorum’ in four
[...] Read more.
The distribution of ‘Candidatus Phytoplasma prunorum’, a phloem-limited pathogen, varies among plant tissues during the year, but knowledge about quantitative comparisons of its presence in roots and above-ground tissues across Prunus species remains limited. The variation in ‘Ca. P. prunorum’ in four Prunus species was monitored monthly during 2025 using quantitative PCR-based absolute quantification. Roots, one-year-old shoots and annual shoots were sampled monthly from 21 infected trees of Prunus armeniaca, Prunus domestica, Prunus persica and Prunus salicina. Phytoplasma titer varied with plant part and sampling month, and differences were also observed among the examined host groups. Because cultivar and rootstock were linked to species, these differences cannot be attributed to species alone. Roots provided the most consistent year-round detection, whereas above-ground tissues showed stronger monthly variation during the studied period. P. domestica displayed the most distinct pattern: roots remained frequently positive, while one-year-old shoots and particularly annual shoots had lower detection rates and lower phytoplasma titers. Multilocus genotyping detected ten multilocus profiles, but their uneven distribution among only 21 trees precluded a reliable evaluation of their relationship with phytoplasma titer or symptom expression.
Full article
(This article belongs to the Special Issue Phytoplasmas and Phytoplasma Diseases)
►▼
Show Figures

Figure 1
Open AccessArticle
Multicentre Evaluation of Cepheid Xpert® Research-Use-Only Panel for Molecular Rapid Detection of Gastrointestinal Pathogens
by
Olivier Dauwalder, Jan Kehrmann, Stéphane Corvec, Maroussia Roelens, Tiphaine Roussel-Gaillard, Edoardo Bixio, Jan Buer, Elisa Baillemont, Martin Prodel, Christophe Martinaud and Valeria Cento
Microorganisms 2026, 14(9), 2011; https://doi.org/10.3390/microorganisms14092011 - 10 Sep 2026
Abstract
Rapid and accurate identification of enteric pathogens is essential for the management of acute infectious diarrhoeal diseases. Multiplex PCR panels have emerged as first-line screening tools to conventional stool culture by providing broad pathogen coverage and shorter turnaround times. This study evaluated the
[...] Read more.
Rapid and accurate identification of enteric pathogens is essential for the management of acute infectious diarrhoeal diseases. Multiplex PCR panels have emerged as first-line screening tools to conventional stool culture by providing broad pathogen coverage and shorter turnaround times. This study evaluated the diagnostic performance of the Cepheid Xpert gastrointestinal (GI) research-use-only (RUO) panel in routine clinical practice across multiple European centres. A multicentre study was conducted in four hospitals across France, Germany and Italy. Stool specimens collected as part of routine care were tested using local standard-of-care (SoC) methods, including conventional stool culture, BDMax® Enteric Panels, or BioFire FilmArray® GI Panel, and subsequently analysed using the Cepheid Xpert GI RUO panel. A total of 11 bacterial, viral and parasitic enteropathogens were targeted. Discordant results underwent additional testing, and a consensus result based on a two-out-of-three agreement rule was used as the reference standard. Diagnostic performance metrics, agreement with SoC methods, microbiological findings and turnaround times were assessed. Of 508 samples, 115 (22.6%) were identified as positive for at least one pathogen according to the consensus result, including 12 samples with co-detection of more than one pathogen (10.4% of positive samples), resulting in a total of 128 pathogens detected. The most frequently detected pathogens were Campylobacter spp. (37 specimens), Norovirus (26 specimens), and Salmonella spp. (25 specimens). Compared with the consensus reference standard, Xpert achieved an overall sensitivity of 98% (95% CI: 94–100%), specificity of 100% (95% CI: 100–100%), positive predictive value of 95% (95% CI: 90–98%), and negative predictive value of 100% (95% CI: 100–100%). Sensitivity was ≥95% and specificity ≥99% for all evaluable targets. Agreement with routine diagnostic methods was high, with overall positive and negative percent agreements exceeding 90% and a Cohen’s kappa coefficient of 0.91. Median analytical turnaround time was 1 h 18 min for Xpert, compared with 3 h for BDMax, 1 h 15 min for BioFire and 48 h for conventional stool culture. The Cepheid Xpert GI (RUO) panel provided highly accurate detection of enteric pathogens across heterogeneous European diagnostic settings and reduced turnaround time by more than 95% compared with conventional stool culture. A future IVD version of this test could provide rapid and accurate detection of enteric pathogens for potential routine management of patients with suspected acute infectious gastroenteritis.
Full article
(This article belongs to the Section Gut Microbiota)
►▼
Show Figures

Figure 1
Open AccessArticle
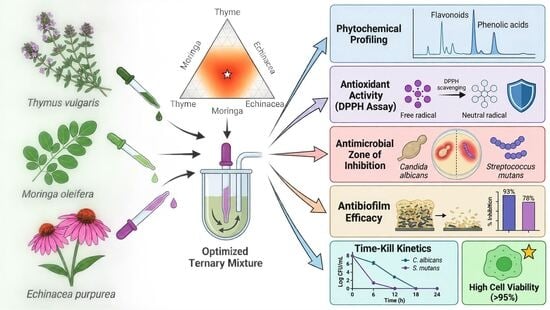
Synergistic Antimicrobial and Antibiofilm Activity Optimization of a Thymus vulgaris–Moringa oleifera–Echinacea purpurea Ternary Ethanolic Extract Blend Against Candida albicans and Streptococcus mutans Using L-Optimal Mixture Design
by
Khadijah A. Altammar
Microorganisms 2026, 14(9), 2010; https://doi.org/10.3390/microorganisms14092010 - 10 Sep 2026
Abstract
Antimicrobial resistance and biofilm-associated oral infections require multi-target therapeutic strategies beyond single-extract herbal testing. Using the L-optimal mixture design approach, we optimized a ternary combination of Thymus vulgaris, Moringa oleifera, and Echinacea purpurea ethanolic extracts against Candida albicans and Streptococcus mutans
[...] Read more.
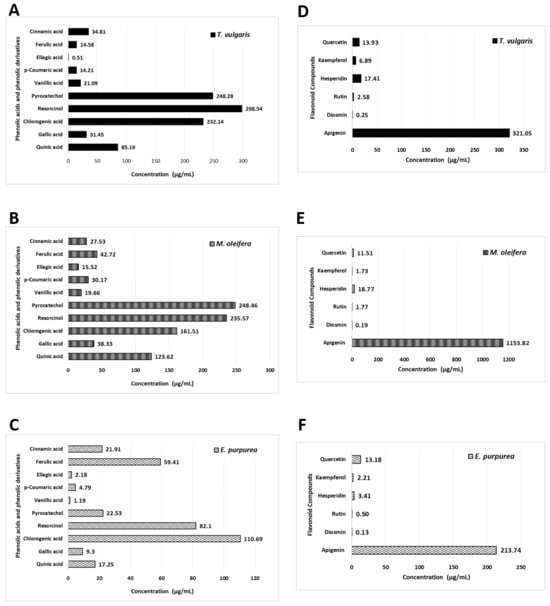
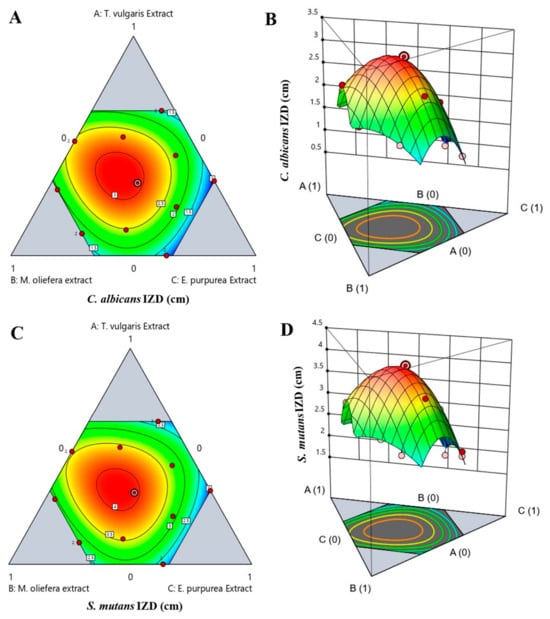
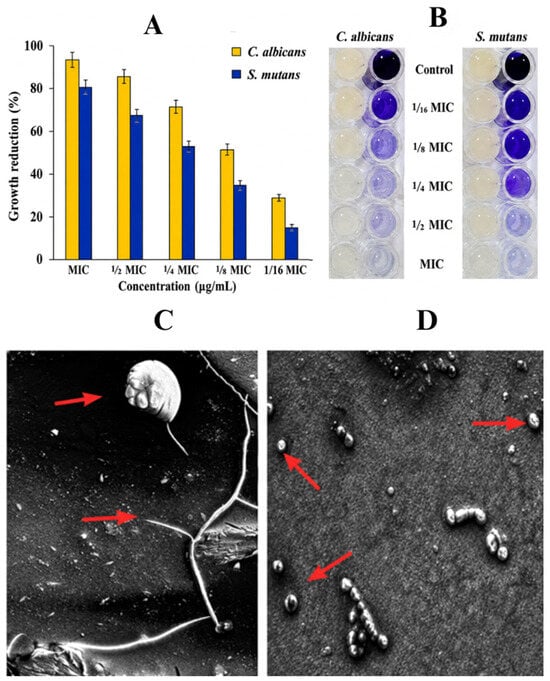
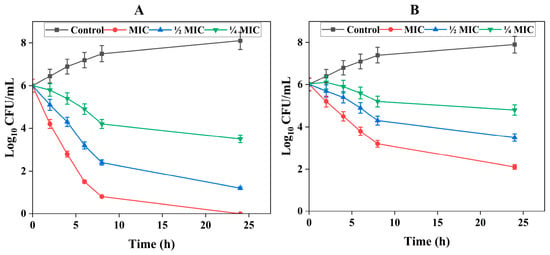
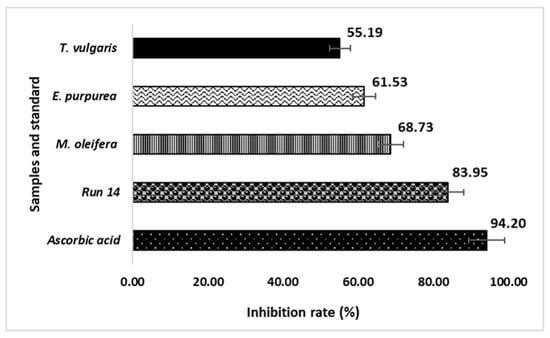
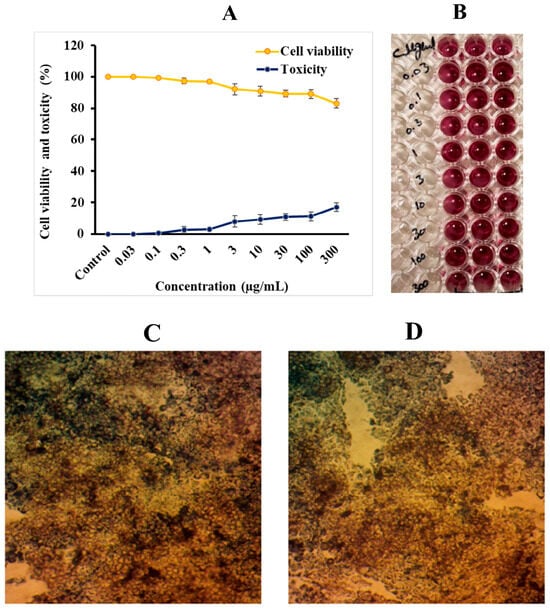
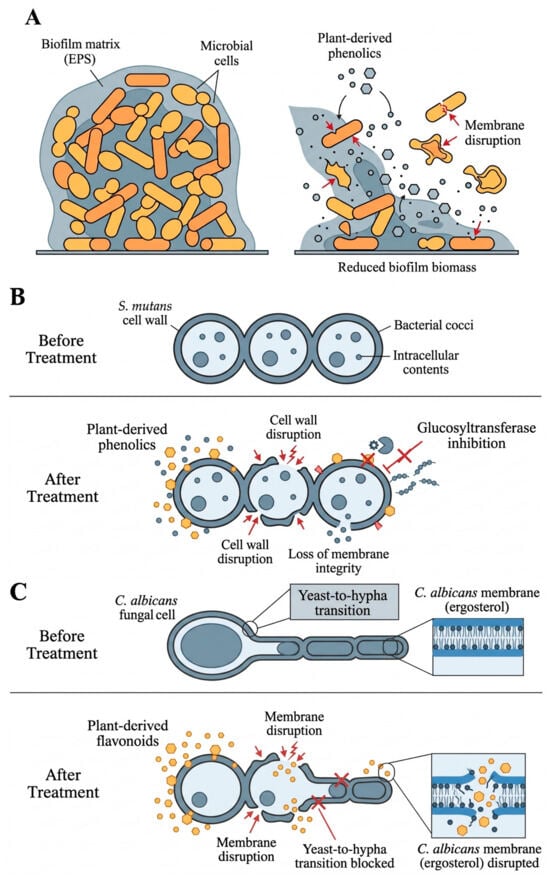
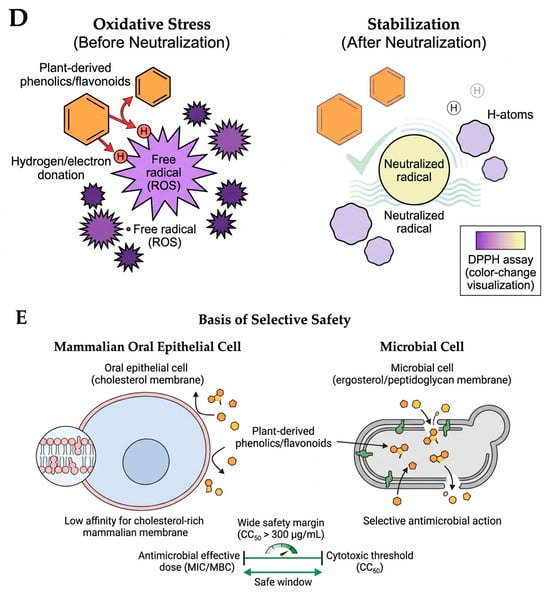
Antimicrobial resistance and biofilm-associated oral infections require multi-target therapeutic strategies beyond single-extract herbal testing. Using the L-optimal mixture design approach, we optimized a ternary combination of Thymus vulgaris, Moringa oleifera, and Echinacea purpurea ethanolic extracts against Candida albicans and Streptococcus mutans, then characterized the optimized blend’s phytochemistry, antimicrobial and antibiofilm efficacy, time–kill pharmacodynamics, antioxidant capacity, and cytotoxic safety. Through HPLC, apigenin, chlorogenic acid, resorcinol, and ferulic acid were identified as dominant constituents. The optimal blend (Run 14: T. vulgaris 0.331, M. oleifera 0.318, E. purpurea 0.352) achieved a fractional inhibitory concentration index of 0.50 against both organisms, maximum inhibition zones of 3.10 cm (C. albicans) and 4.10 cm (S. mutans), biofilm inhibition of up to 93.2 ± 2.1% for C. albicans and 78.4 ± 2.8% for S. mutans, and confirmed fungicidal and bactericidal activity within 48 and 24 h, respectively. DPPH scavenging ranged from 55.19 to 68.73%, and oral epithelial cells (OEC) viability exceeded 83% at 300 µg/mL. All assays included three independent biological replicates; statistical significance was defined as p < 0.05. These findings identify the T. vulgaris–M. oleifera–E. purpurea blend as a synergistic, multi-target antimicrobial candidate with a favorable preliminary safety profile, supporting further development and evaluation against clinical isolates for oral fungal and bacterial infections.
Full article
(This article belongs to the Section Antimicrobial Agents and Resistance)
►▼
Show Figures

Graphical abstract
Open AccessArticle
Integrating Microbial Indicators from High-Throughput Sequencing into Soil Quality Index: A Case Study in a Restored Mining Area
by
Zhengjun Feng, Chaolong Ma, Shengxin Yan, Wenhui Liu, Peiyin Li, Yan Zou, Dashdorj Munkhbat and Huiping Song
Microorganisms 2026, 14(9), 2009; https://doi.org/10.3390/microorganisms14092009 - 10 Sep 2026
Abstract
The Soil Quality Index (SQI) is a vital tool for evaluating soil quality; however, traditional approaches seldom integrate microbial data from high-throughput sequencing—commonly used to characterize soil microbial communities—into the SQI framework. This study enhances the SQI by incorporating microbial indicators derived from
[...] Read more.
The Soil Quality Index (SQI) is a vital tool for evaluating soil quality; however, traditional approaches seldom integrate microbial data from high-throughput sequencing—commonly used to characterize soil microbial communities—into the SQI framework. This study enhances the SQI by incorporating microbial indicators derived from high-throughput sequencing, establishing a more comprehensive evaluation system. We collected soil samples from mining areas and analyzed their fundamental physicochemical properties and microbial indicators. Four SQI models were constructed using different indicator sets: (1) only physicochemical properties (T-SQI); (2) physicochemical properties and bacterial α-diversity (α-SQI); (3) physicochemical properties, α-diversity, and relative abundances of the top five abundant bacteria (αMA-SQI); and (4) physicochemical properties, α-diversity, and relative abundances of the top five bacteria based on LDA scores (αBM-SQI). Results demonstrated that integrating multi-level microbial indicators improved the rationality of soil quality rankings and significantly strengthened correlations with α-diversity. Gemmatimonadota was consistently selected in the Minimum Data Set (MDS) for both αMA-SQI and αBM-SQI, highlighting its ecological importance. Statistically, microbial indicators at the order and family levels were most suitable for inclusion in the MDS, as their results deviated least from the total dataset. In conclusion, incorporating microbial diversity across taxonomic levels refines the SQI, enabling a more accurate and holistic assessment of soil health.
Full article
(This article belongs to the Section Environmental Microbiology)
►▼
Show Figures

Figure 1
Open AccessArticle
Lactococcus G423 Improves Lipid Metabolism in Broilers Through the Gut Microbiota and AMPK Signaling Pathway
by
Bo Pang, Ruixiang Li, Xinyu Wang, Weitong Guan, Wei Ma, Chunqiang Wang, Desheng Li and Mi Wang
Microorganisms 2026, 14(9), 2008; https://doi.org/10.3390/microorganisms14092008 - 10 Sep 2026
Abstract
This study aimed to elucidate the role of Lactococcus G423 in regulating the AMPK (AMP-activated protein kinase) signaling pathway and gut microbiota, and to investigate the mechanisms underlying its potential to improve lipid metabolism in broilers. In this trial, 960 one-day-old Arbor Acres
[...] Read more.
This study aimed to elucidate the role of Lactococcus G423 in regulating the AMPK (AMP-activated protein kinase) signaling pathway and gut microbiota, and to investigate the mechanisms underlying its potential to improve lipid metabolism in broilers. In this trial, 960 one-day-old Arbor Acres (AA) broilers were randomly assigned to one of three groups: a control group (CON), a low-dose Lactococcus G423 group (Lac-L), and a high-dose Lactococcus G423 group (Lac-H). Each group consisted of eight replicates, with 40 individuals per replicate. Compared with the CON group, the Lac-L and Lac-H groups showed no significant differences in ADFI (Average Daily Feed Intake), ADG (Average Daily Gain), FCR (Feed Conversion Ratio), breast muscle yield, leg muscle yield, or abdominal fat yield (p > 0.05). Compared with the control group, neither Lac-L nor Lac-H affected serum TG (Triglyceride) or LDL (Low-Density Lipoprotein) levels (p > 0.05). However, both Lac-L and Lac-H significantly decreased serum CHO (Cholesterol) and increased serum HDL (High-Density Lipoprotein) (p < 0.05). Regarding the AMPK signaling pathway, both the Lac-L and Lac-H groups significantly downregulated the hepatic mRNA expression of Acetyl-CoA carboxylase (ACC) (p < 0.05). The Lac-L groups significantly increased the mRNA expression of AMP-activated protein kinase (AMPK) and Sterol Regulatory Element-Binding Protein 1 (SREBP1) (p < 0.05), and the Lac-H groups significantly increased the mRNA expression of Apolipoprotein A1 (APOA1) and Protein kinase B (PKB) (p < 0.05) and significantly reduced the mRNA expression of Fatty acid-binding protein 1 (FABP1) (p < 0.05), compared to the CON group. Gut microbiota analysis indicated that both treatment groups exhibited significantly increased α-diversity indices (p < 0.05) and altered microbial composition at an order level comparable with that of the CON group. Specifically, the abundances of beneficial families such as Lachnospirales and Lactobacillales were elevated, while significant changes were observed in other taxa including Ruminococcaceae and Akkermansiaceae (p < 0.05). These changes corresponded with significant differences in the MDI and GMHI (p < 0.05). In conclusion, Lactococcus G423 may regulate lipid metabolism in broilers through the combined modulation of the AMPK signaling pathway and the gut microbiota. Specifically, Lactococcus G423 appears to regulate broiler lipid metabolism by modulating the gut microbiota–hepatic lipid metabolism axis.
Full article
(This article belongs to the Section Food Microbiology)
►▼
Show Figures

Figure 1
Open AccessArticle
Th1 and Th17 Responses to LTB and Colonization Factors Following Oral ETEC Vaccination
by
Joanna Kaim and Anna Lundgren
Microorganisms 2026, 14(9), 2007; https://doi.org/10.3390/microorganisms14092007 - 10 Sep 2026
Abstract
T helper cells (Th) are central to mucosal IgA induction and key targets for modulation by vaccine adjuvants. To improve understanding of cellular mechanisms underlying mucosal vaccine-induced immunity, we analyzed antigen-specific peripheral blood Th responses elicited by the oral enterotoxigenic Escherichia coli (ETEC)
[...] Read more.
T helper cells (Th) are central to mucosal IgA induction and key targets for modulation by vaccine adjuvants. To improve understanding of cellular mechanisms underlying mucosal vaccine-induced immunity, we analyzed antigen-specific peripheral blood Th responses elicited by the oral enterotoxigenic Escherichia coli (ETEC) vaccine ETVAX, administered with or without the double mutant heat-labile toxin (dmLT) adjuvant. ETVAX, consisting of inactivated E. coli overexpressing colonization factors CFA/I, CS3, CS5, and CS6 with a heat-labile toxin B-subunit toxoid, was given orally in two doses to adult volunteers, either alone or with 10 or 25 µg dmLT. Antigen-specific Th-associated cytokine responses were assessed in stimulated peripheral blood mononuclear cells isolated from 15 to 18 individuals/group using ELISA and electrochemiluminescence assays. ETVAX predominantly induced Th1 (IFN-γ) and Th17 (IL-17A) responses, with minimal Th2-associated cytokines. Responses were markedly reduced after CD4+ T-cell depletion, supporting a Th cell origin. The strongest responses targeted LTB and CS3, with IFN-γ responses detected in 60–80% and IL-17A in 40–60% across all vaccinees. Responses to CFA/I, CS5 and CS6 were generally weaker. Exploratory comparisons suggested broader IFN-γ responses and more consistent IFN-γ and IL-17A responses to lower-dose antigens, particularly CS6, in recipients receiving vaccine plus 10 µg dmLT. These trends paralleled IgA antibody-secreting cell response patterns, with significantly enhanced IgA responses to CS6 in the vaccine plus 10 µg dmLT group. In conclusion, ETVAX induces antigen-specific Th1- and Th17-type responses in peripheral blood, supporting a role for cellular immunity in mucosal responses to oral ETEC vaccines.
Full article
(This article belongs to the Special Issue Advancement in Enterotoxigenic Escherichia coli (ETEC) Vaccines)
►▼
Show Figures

Figure 1
Open AccessArticle
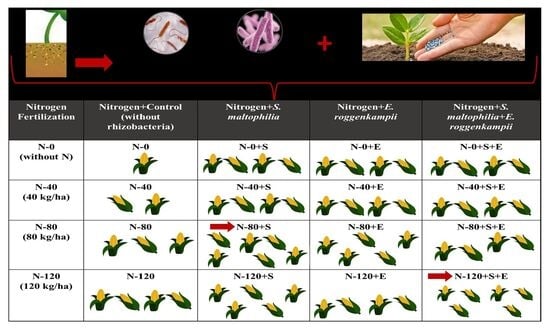
Inoculation with Stenotrophomonas maltophilia LIMN and Enterobacter roggenkampii LCMG Enhances Maize (Zea mays L.) Yield and Reduces Nitrogen Fertilizer Dependence in Nutrient-Limited Soils of Semi-Arid Regions
by
Odilón Gayosso Barragán, Griselda Chávez Aguilar, Deli Nazmín Tirado González, Roberto Reynoso Santos, Ismael Fernando Chávez Díaz, Lily Xochilt Zelaya-Molina, Gustavo Tirado Estrada and Luis Yobani Gayosso Rosales
Microorganisms 2026, 14(9), 2006; https://doi.org/10.3390/microorganisms14092006 - 10 Sep 2026
Abstract
Excessive nitrogen (N) fertilization in maize (Zea mays L.) production causes environmental degradation, particularly in semi-arid regions where high chemical inputs are necessary to maintain crop production. Plant growth-promoting rhizobacteria (PGPR) could reduce the optimal N fertilization doses. This investigation aimed to
[...] Read more.
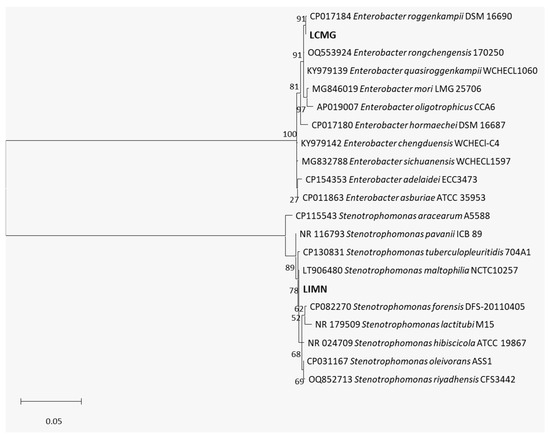
Excessive nitrogen (N) fertilization in maize (Zea mays L.) production causes environmental degradation, particularly in semi-arid regions where high chemical inputs are necessary to maintain crop production. Plant growth-promoting rhizobacteria (PGPR) could reduce the optimal N fertilization doses. This investigation aimed to isolate bacteria, screen them for plant growth-promoting traits, and evaluate their potential effects on maize grain under different chemical N fertilization doses. Isolates Stenotrophomonas maltophilia LIMN and Enterobacter roggenkampii LCMG demonstrated multifunctional growth-promoting traits in vitro, including N2 fixation, indole-3-acetic acid and siderophore production, potassium solubilization, and desiccation tolerance, with E. roggenkampii LCMG also solubilizing inorganic phosphate. A field trial across four chemical N fertilization rates (0, 40, 80, and 120 kg N ha−1) revealed that single bacterial inoculation without N fertilizer matched or exceeded uninoculated controls receiving up to 80 kg of N ha−1 (the standard recommendation). Specifically, S. maltophilia LIMN combined with the recommended rate of 80 kg N ha−1 achieved the maximum grain yield (4601.2 kg DM ha−1), outperforming uninoculated controls (without PGPR) receiving excessive fertilization at 120 kg N ha−1 (3760.4 DM ha−1). Although maize plants benefited from inoculation with S. maltophilia LIMN or E. roggenkampii LCMG, achieving similar or better yields than crops fertilized with high chemical N inputs under stress factors such as deficiency of moisture, low-nutrient soils, and high temperatures, the present study did not test the pathogenicity and biosecurity of S. maltophilia or E. roggenkampii in crops; therefore, the results are not a direct recommendation of their use as biofertilizers before novel studies to assess all the limitations and the potential to reduce N dependency in order to design novel sustainable strategies for crop production.
Full article
(This article belongs to the Topic Applications of Biotechnology in Food and Agriculture)
►▼
Show Figures

Graphical abstract
Open AccessEditorial
Editorial for the Special Issue “Advances in Microbial and Plant Biotechnology”
by
Svetlana Veselova and Igor Maksimov
Microorganisms 2026, 14(9), 2005; https://doi.org/10.3390/microorganisms14092005 - 10 Sep 2026
Abstract
Introduction [...]
Full article
(This article belongs to the Special Issue Advances in Microbial and Plant Biotechnology)
Open AccessArticle
Antimicrobial Resistance Profiles, Resistance Genes, and Their Association with Bacteriophage Susceptibility Among Vibrio parahaemolyticus Isolates from Zhanjiang Shrimp Aquaculture Farms
by
Lukman Iddrisu, Baochen Chai, Evodia Moses Mkulo, Salifu Ibrahim, Felix Danso, Muqadas Altaf, Muhammad Fahad Khan, Bingyu Zhang, Shumei Zhang, Jiesen Su, Yinyan Chen, Zhijia Fang and Jianzhi Ye
Microorganisms 2026, 14(9), 2004; https://doi.org/10.3390/microorganisms14092004 - 9 Sep 2026
Abstract
Vibrio parahaemolyticus is an important foodborne and aquaculture-associated pathogen, and its increasing antimicrobial resistance threatens shrimp production, seafood safety, and public health. Although bacteriophages have emerged as a potential solution to the growing challenge of antimicrobial resistance, whether antibiotic resistance is associated with
[...] Read more.
Vibrio parahaemolyticus is an important foodborne and aquaculture-associated pathogen, and its increasing antimicrobial resistance threatens shrimp production, seafood safety, and public health. Although bacteriophages have emerged as a potential solution to the growing challenge of antimicrobial resistance, whether antibiotic resistance is associated with phage tolerance remains unclear. This study characterized antimicrobial resistance profiles and resistance genes in Vibrio parahaemolyticus isolates from Zhanjiang shrimp aquaculture farms and evaluated their association with bacteriophage susceptibility. A total of 132 isolates were tested against 18 antibiotics, while molecular and phage-related analyses were performed on 132 isolates. High resistance was observed against kanamycin (95.5%), sulfamethoxazole (94.7%), cefazolin (78.8%), imipenem (78.8%), amoxicillin (70.5%), and ampicillin (65.2%). Multidrug resistance was detected in 131/132 isolates (99.24%), indicating an extensive resistance burden. PCR analysis showed high detection rates of aphA (100.00%), sul1 (99.24%), blaTEM (98.48%), blaOXA (73.48%), blaCTX-M (53.03%), qnrA (45.45%), tetA (37.88%), and tetB (21.21%). Phage response analysis showed that 70.45% and 68.18% of isolates were phage-resistant at 4 h and 5 h, respectively. Genes potentially associated with phage–host interactions were also detected, with luxS showing the highest frequency (85.61%). Association analysis revealed significant links between tetA, tetB, blaOXA, and bacteriophage responses. These findings suggest that antimicrobial resistance and reduced phage susceptibility can co-occur among V. parahaemolyticus isolates recovered from shrimp farms in Zhanjiang, highlighting the need for regional resistance surveillance and careful phage selection.
Full article
(This article belongs to the Special Issue Foodborne Pathogens: Detection, Resistance, Risk Assessment and Control)
►▼
Show Figures

Figure 1
Open AccessArticle
Whole-Genome Sequencing of Escherichia coli O18ab:H11 from South African Beef: Antimicrobial Resistance, Virulence, and a Rare Sequence Type
by
Fezeka P. Ndlazi, Ephifania Geza and Evelyn Madoroba
Microorganisms 2026, 14(9), 2003; https://doi.org/10.3390/microorganisms14092003 - 9 Sep 2026
Abstract
Antimicrobial-resistant Escherichia coli in food systems represents a growing public health concern, yet whole-genome sequencing (WGS)-based surveillance of beef-associated E. coli in South Africa remains limited. This study applied WGS to comprehensively characterise the antimicrobial resistance (AMR) profile, virulence gene repertoire, sequence type,
[...] Read more.
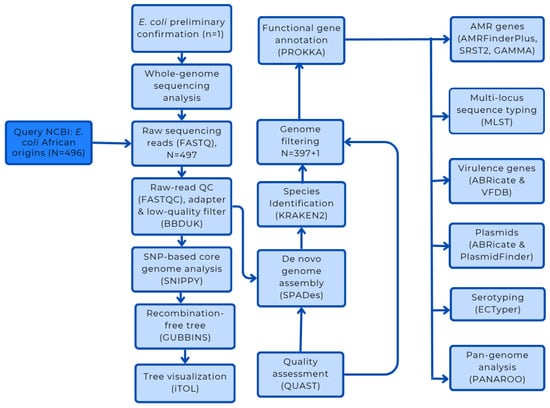
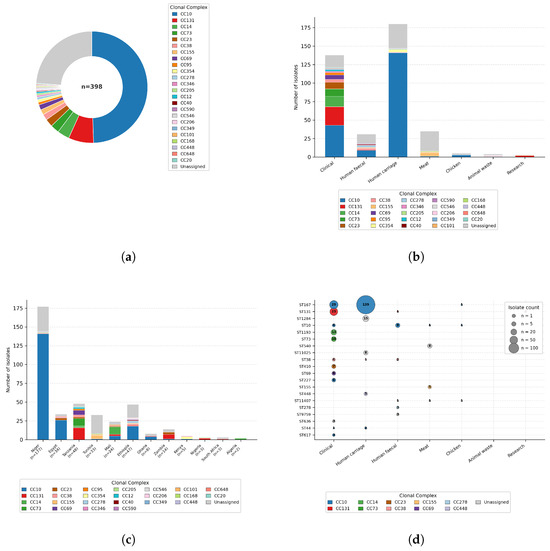
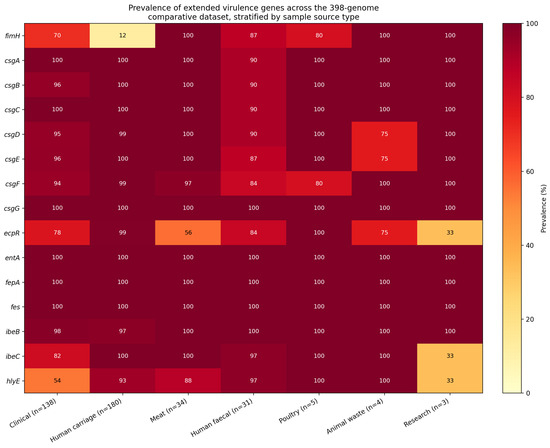
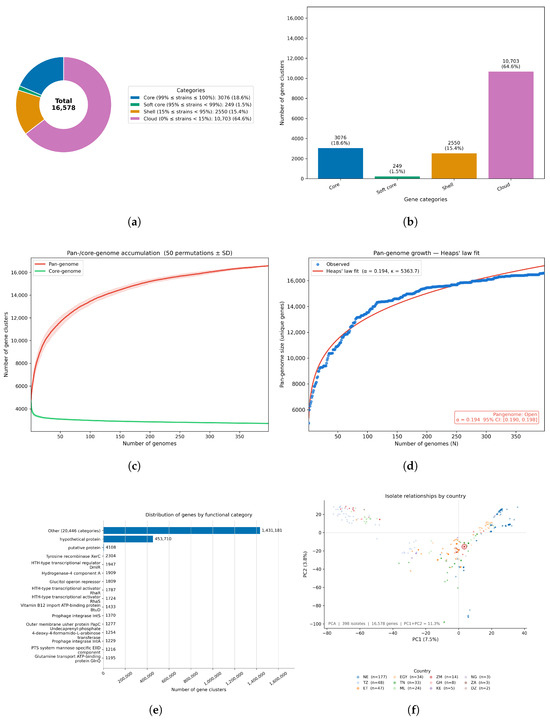
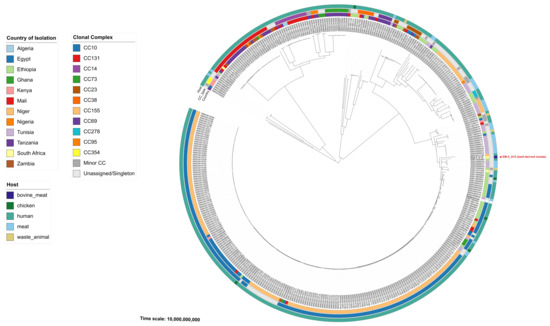
Antimicrobial-resistant Escherichia coli in food systems represents a growing public health concern, yet whole-genome sequencing (WGS)-based surveillance of beef-associated E. coli in South Africa remains limited. This study applied WGS to comprehensively characterise the antimicrobial resistance (AMR) profile, virulence gene repertoire, sequence type, and serotype of an E. coli isolate recovered from retail beef in KwaZulu-Natal Province, South Africa, and to contextualise these findings within the broader African E. coli population through comparative genomic analysis of 397 publicly available E. coli genome sequences retrieved from the NCBI SRA. The isolate was recovered by classical microbiological methods, confirmed by MALDI-TOF MS, and whole-genome-sequenced on the PacBio Onso platform. In silico serotyping assigned the genome to serotype O18ab:H11. MLST under the Achtman seven-locus scheme assigned the genome to ST7106, a sequence type with no prior record in South Africa. The isolate carried the intrinsic -lactamase gene blaEC-15 (reported as chromosomal in the literature), conferring narrow-spectrum resistance to aminopenicillins, alongside an acquired resistome comprising aadA1, tet(B), and sat2, conferring resistance to aminoglycosides, tetracyclines, and streptothricin; no ESBL or carbapenemase genes were detected. The virulence gene repertoire—encompassing type 1 fimbriae (fimA–I), curli fibres (csgA–G), the E. coli common pilus (ecpRABCDE), enterobactin iron-acquisition genes, hlyE, and the invasion genes ibeB and ibeC—indicates colonisation capacity, but ibeB, ibeC, and hlyE showed no clinical enrichment relative to carriage genomes, and the isolate showed no close genomic relationships with any clinical genome; a definitive extra-intestinal pathogenic E. coli (ExPEC) pathotype assignment is therefore not supported. PathogenFinder returned a pathogenicity probability of 0.944 (532 pathogen-associated protein families detected), without in vivo corroboration. Comparative analysis identified a pan-resistome of 97 unique AMR genes across the 398 genomes, including blaCTX-M-15 (59.0%) and blaNDM-5 (49.5%), predominantly among clinical genomes. Pan-genome analysis revealed an open genomic structure ( ; 16,578 gene clusters), and core-genome phylogenetics confirmed the beef-derived isolate as a genetically distinct lineage (minimum pairwise SNP distance: 1629 SNPs). These findings demonstrate the presence of a genomically distinct, potentially pathogenic E. coli lineage in the South African beef supply and highlight the importance of integrating WGS into routine food safety and AMR surveillance frameworks across the region.
Full article
(This article belongs to the Section Food Microbiology)

►▼
Show Figures

Figure 1
Open AccessArticle
High-Throughput Sequencing Reveals Composition, Diversity, and Functional Prediction of Root-Associated Microbial Communities of Dominant Plants in an Ecologically Sensitive Area of the Loess Plateau
by
Gexue Bai, Qingqing Tan, Bingbing Han, Ruidong Li, Lijun Gu, Xiaojing Wang, Jie Zhang, Yan Li and Quanfang Zhang
Microorganisms 2026, 14(9), 2002; https://doi.org/10.3390/microorganisms14092002 - 9 Sep 2026
Abstract
The alpine mining area of the Qilian Mountains features a fragile ecosystem, severe soil degradation due to mining disturbances, and slow natural recovery. To clarify the ecological restoration potential of rhizosphere microorganisms associated with native dominant plants, this study investigated the community structure,
[...] Read more.
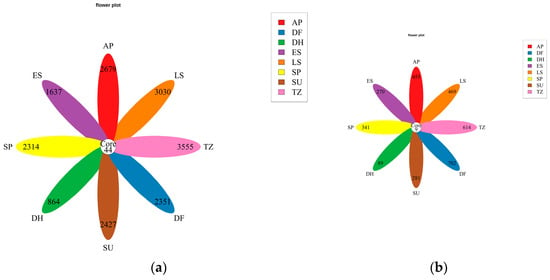
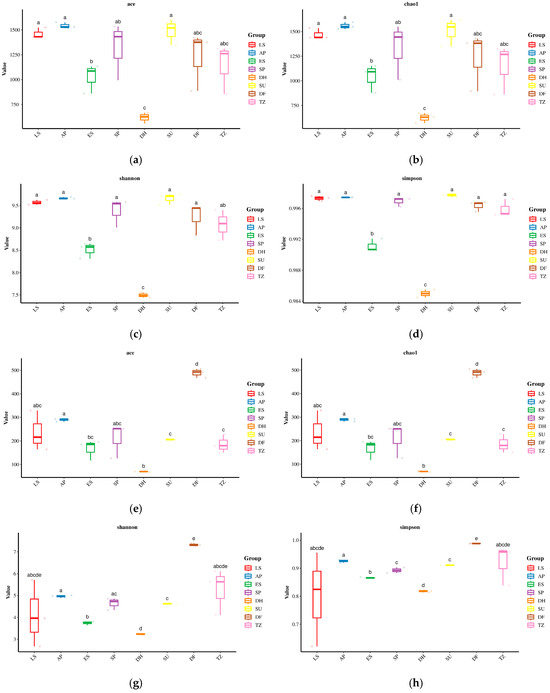
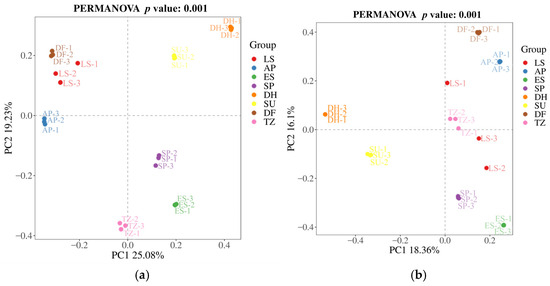
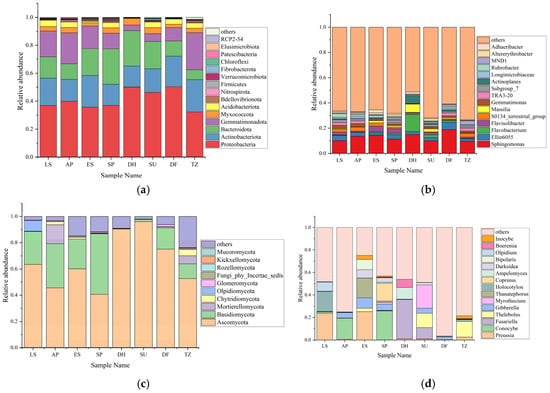
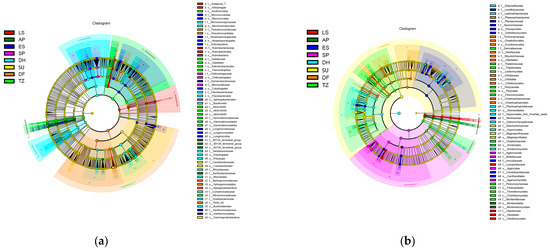
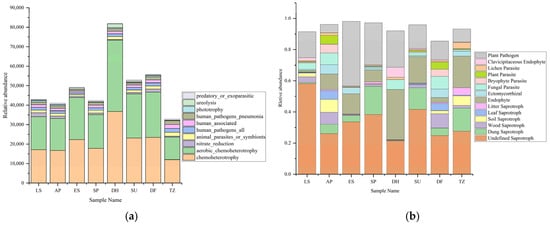
The alpine mining area of the Qilian Mountains features a fragile ecosystem, severe soil degradation due to mining disturbances, and slow natural recovery. To clarify the ecological restoration potential of rhizosphere microorganisms associated with native dominant plants, this study investigated the community structure, diversity, and functional differentiation patterns of rhizosphere bacterial and fungal communities associated with seven native dominant plant species in the Tianzhu mining area. Using Illumina NovaSeq 6000 high-throughput sequencing, we amplified the bacterial 16S rRNA V3–V4 region and the fungal ITS1 region. A total of 24 sequencing libraries (seven plant species + bare soil, three replicates each) were analyzed. Bioinformatics analyses examined ASV distributions, alpha/beta diversity, species composition, and differentially enriched taxa, followed by functional predictions. Bacterial alpha diversity (Chao1: 623.39–1553.25; Shannon: 7.49–9.67) varied significantly among plant species, with Allium przewalskianum (AP) showing the highest bacterial richness and diversity (Chao1 = 1553.25 ± 35.89, Shannon = 9.67 ± 0.02). Fungal alpha diversity also showed significant variation (Chao1: 69.66–488.56; Shannon: 3.24–4.96), with Dasiphora fruticosa (DF) exhibiting the highest fungal diversity (Chao1 = 488.56 ± 18.71, Shannon = 7.32 ± 0.06). At the phylum level, Proteobacteria (32.22–50.32%) and Actinobacteriota (15.04–23.19%) were core bacterial groups, and Ascomycota (45.54–96.07%) dominated fungal communities. PERMANOVA confirmed significant differences in community composition among plant species (bacteria: R2 = 0.78, p < 0.001; fungi: R2 = 0.84, p < 0.001). Different plant species were associated with distinct predicted functional taxa, which may serve as candidate biomarkers for soil remediation. However, all functional interpretations are predictive and require experimental validation. In conclusion, rhizosphere microbial community composition and predicted functional profiles differed among native plant species, providing correlative evidence and candidate targets for future vegetation–microbe synergy studies in alpine mining area restoration.
Full article
(This article belongs to the Special Issue Microbial Interactions and Community Assembly Mechanisms)
►▼
Show Figures

Figure 1
Open AccessReview
Ecological Engineering of the Human Gut Microbiome: A Narrative Review and Framework for Next-Generation Therapeutics
by
Antonio Díaz, Gissel García and Raúl De Jesús Cano
Microorganisms 2026, 14(9), 2001; https://doi.org/10.3390/microorganisms14092001 - 9 Sep 2026
Abstract
The human gut microbiome is a complex adaptive ecosystem whose functions arise from interactions among microbial populations rather than from isolated taxa. Nevertheless, many microbiome-directed interventions still rely on administering individual strains, with limited consideration of the ecological processes governing community assembly, succession,
[...] Read more.
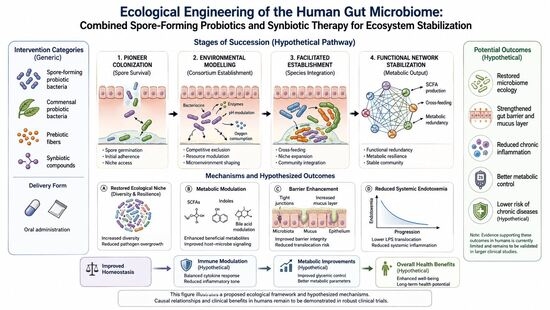
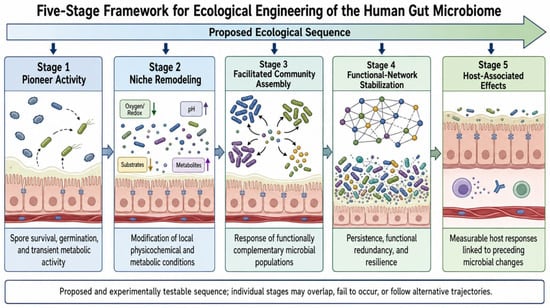
The human gut microbiome is a complex adaptive ecosystem whose functions arise from interactions among microbial populations rather than from isolated taxa. Nevertheless, many microbiome-directed interventions still rely on administering individual strains, with limited consideration of the ecological processes governing community assembly, succession, and resilience. This review integrates evidence from microbial ecology, comparative genomics, systems biology, mechanistic physiology, and clinical microbiome research to propose a testable framework for ecologically engineering the human gut microbiome. Within this framework, selected spore-forming probiotics are hypothesized to function as transient pioneer organisms that modify intestinal physicochemical and metabolic conditions, thus facilitating the establishment and activity of functionally complementary microbial populations delivered through rationally designed synbiotic consortia. The proposed process comprises five stages: pioneer activity, niche remodeling, facilitated community assembly, functional-network stabilization, and the emergence of host-associated outcomes. Available genomic, physiological, and clinical observations support the biological plausibility of individual components of this model but do not yet demonstrate directed ecological succession as a complete causal process. Accordingly, the framework distinguishes established evidence from ecological inference and generates experimentally testable predictions of temporal niche modification, metabolic cross-feeding, functional redundancy, resilience after treatment withdrawal, and host metabolic responses. This ecological perspective shifts the objective of microbiome therapeutics from transient strain supplementation toward the predictable modulation of community trajectories, providing an experimental foundation for developing more resilient, mechanism-based interventions.
Full article
(This article belongs to the Collection Feature Papers in Gut Microbiota Research)
►▼
Show Figures

Graphical abstract
Open AccessArticle
High-Resolution Genomic Profiling of Campylobacter jejuni Across Human, Livestock, and Yellow-Legged Gull Reservoirs in Croatia Reveals Lineage Segregation and Shared Environmental Genotypes
by
Sanja Duvnjak, Luka Jurinović, Fani Krstulović, Viktor Mašović, Andrea Humski, Gordan Kompes, Antonela Bagarić, Eva Postružin, Irena Reil, Merica Carev, Domagoj Drenjančević, Silvija Šoprek Strugar, Arjana Tambić Andrašević, Mirna Vranić-Ladavac, Jasmina Kučinar, Louie Thomas Taylor and Biljana Ječmenica
Microorganisms 2026, 14(9), 2000; https://doi.org/10.3390/microorganisms14092000 - 9 Sep 2026
Abstract
Campylobacter jejuni remains a leading cause of foodborne gastroenteritis worldwide, yet the epidemiological role of wild birds as reservoirs remains poorly understood. This study investigated the genetic diversity, transmission dynamics, and virulome architecture of 396 C. jejuni isolates collected in Croatia (2021–2025) from
[...] Read more.
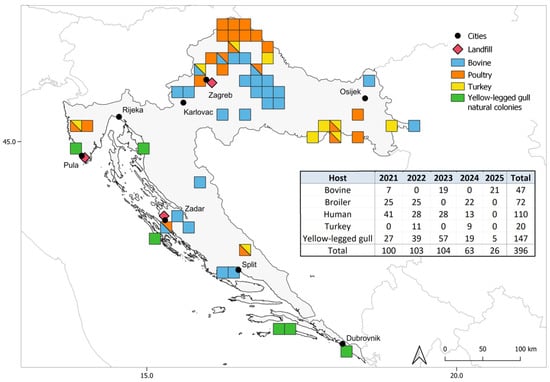
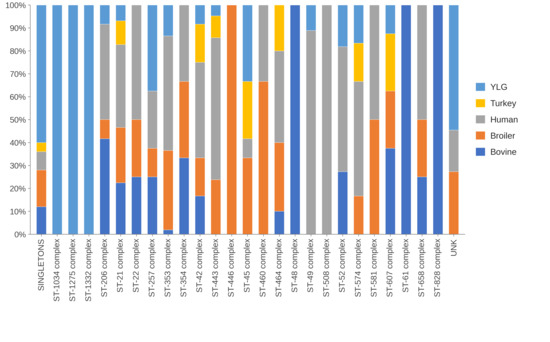
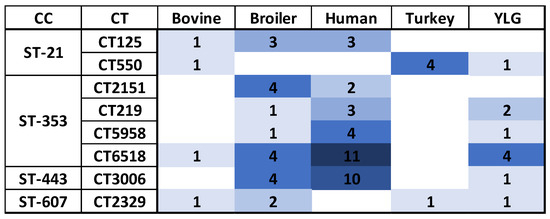
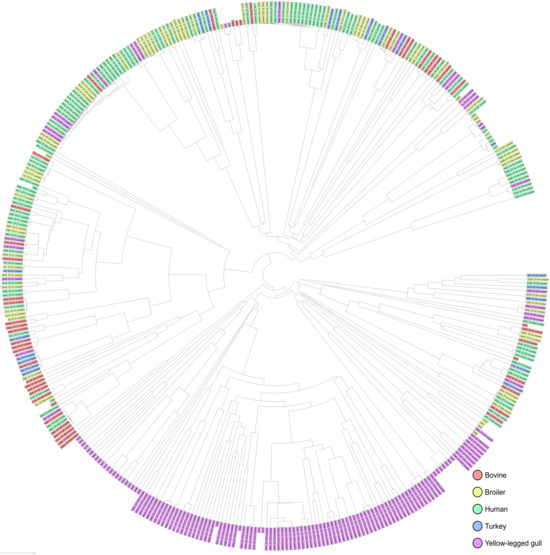
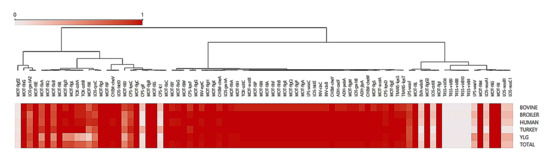
Campylobacter jejuni remains a leading cause of foodborne gastroenteritis worldwide, yet the epidemiological role of wild birds as reservoirs remains poorly understood. This study investigated the genetic diversity, transmission dynamics, and virulome architecture of 396 C. jejuni isolates collected in Croatia (2021–2025) from human, yellow-legged gull, broiler, bovine, and turkey sources. Whole-genome sequencing revealed high genomic diversity ( ), encompassing 124 sequence types across 25 clonal complexes. While classical MLST clustered 17.01% (n = 25/147, 95% CI: 11.83–23.82%) of gull isolates with human-associated lineages, high-resolution cg/wgMLST refined this overlap, segregating the majority of yellow-legged gull strains into a distinct, host-adapted ST-1275 clade (51.02%, n = 75/147). Furthermore, cg/wgMLST identified shared multi-host genotypes, such as CT6518 (ST-353) and CT3006 (ST-443), across wild gulls, domestic livestock, and clinical human cases. Virulome screening revealed a conserved pathogenic core alongside a lineage-specific periphery; the Guillain-Barré-syndrome-associated wlaN gene showed strong restriction to the dominant ST-21 generalist lineage, whereas the Type IV Secretion System was exceptionally rare (<1.5%). These findings demonstrate that, while most yellow-legged gull isolates constitute host-adapted lineages, gulls also harbor overlapping genotypes shared with livestock and clinical human isolates (6.80%, 95% CI: 3.69–12.18%), compatible with shared ecological niches and common contamination sources rather than direct bird-to-human transmission. This highlights their role as environmental sentinels and potential bridge reservoirs within One Health transmission interfaces. This study provides an indispensable baseline for integrated One Health biosecurity and surveillance strategies that bring together poultry production, ecosystem management, and public health infrastructure.
Full article
(This article belongs to the Special Issue Research on Foodborne Pathogens and Disease, 2nd Edition)
►▼
Show Figures

Figure 1
Open AccessArticle
Integrating Metabolic Modeling and Targeted Supplementation for the Rapid Detection of Clostridium tyrobutyricum in Dairy Products
by
Idris Arslan
Microorganisms 2026, 14(9), 1999; https://doi.org/10.3390/microorganisms14091999 - 9 Sep 2026
Abstract
Clostridium tyrobutyricum is a major cause of late blowing defects (LBDs) in cheese, resulting in substantial economic losses. Early detection is critical for maintaining product quality. In this study, we developed a rapid detection approach integrating genome-scale metabolic modeling (GEM) with systematic culture
[...] Read more.
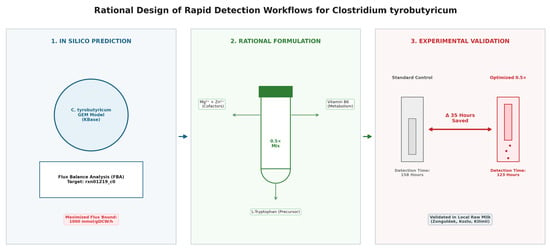
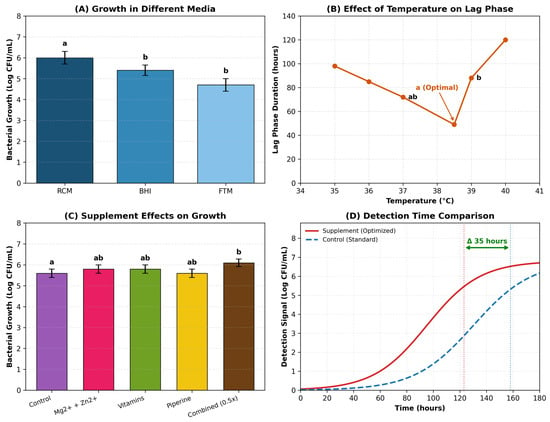
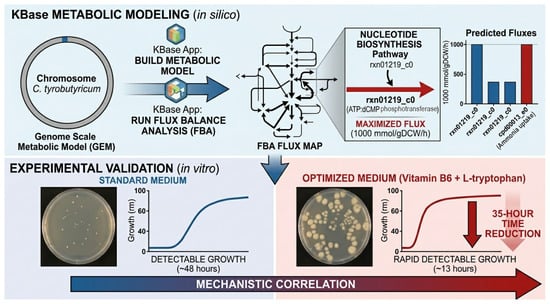
Clostridium tyrobutyricum is a major cause of late blowing defects (LBDs) in cheese, resulting in substantial economic losses. Early detection is critical for maintaining product quality. In this study, we developed a rapid detection approach integrating genome-scale metabolic modeling (GEM) with systematic culture optimization. Three media were evaluated, identifying RCM at 38.5 °C as the optimal condition for reducing the lag phase. Flux Balance Analysis (FBA) revealed that targeted supplementation with magnesium, zinc, Vitamin B6, and L-tryptophan significantly enhanced metabolic flux through nucleotide biosynthesis and energy transfer pathways, particularly reaction rxn01219_c0. Validation using artificially contaminated milk confirmed that the optimized 0.5× supplementation mixture synergistically reduced detection time by approximately 35 h compared to conventional MPN methods. This study demonstrates that bridging systems biology with traditional microbiology provides a cost-effective and mechanistic framework for rapid pathogen detection in the dairy industry.
Full article
(This article belongs to the Section Food Microbiology)
►▼
Show Figures

Graphical abstract
Open AccessArticle
Construction and Comparison of Molecular Identification Methods for Mannheimia haemolytica
by
Zhen Yang, Yongqiang Miao, Hao Mou, Suhui Zhang, Liu Yang, Dengfeng Xu, Lizhi Fu, Kefei Shen, Ziqi Li, Long Zhao and Yuandi Yu
Microorganisms 2026, 14(9), 1998; https://doi.org/10.3390/microorganisms14091998 - 9 Sep 2026
Abstract
Mannheimia haemolytica is a primary pathogen causing bacterial pneumonia in cattle and sheep. Multiple PCR targets exist for its identification, but no standardized primer pairs have been widely adopted. This study evaluated the universality of six reported primers (including rpt2) using clinical
[...] Read more.
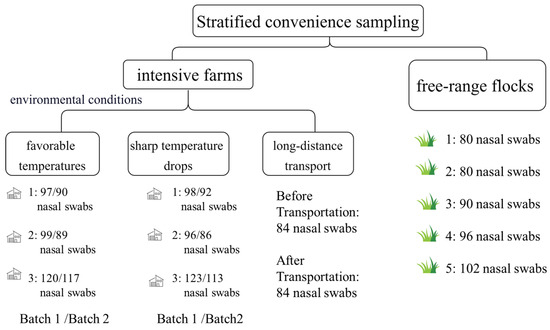
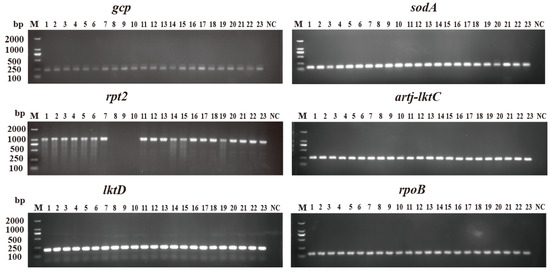
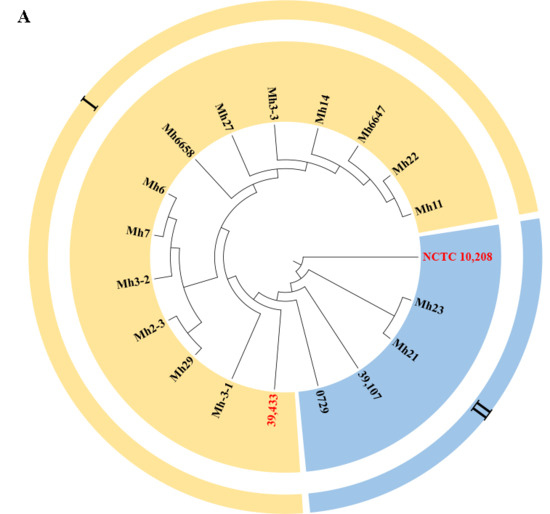
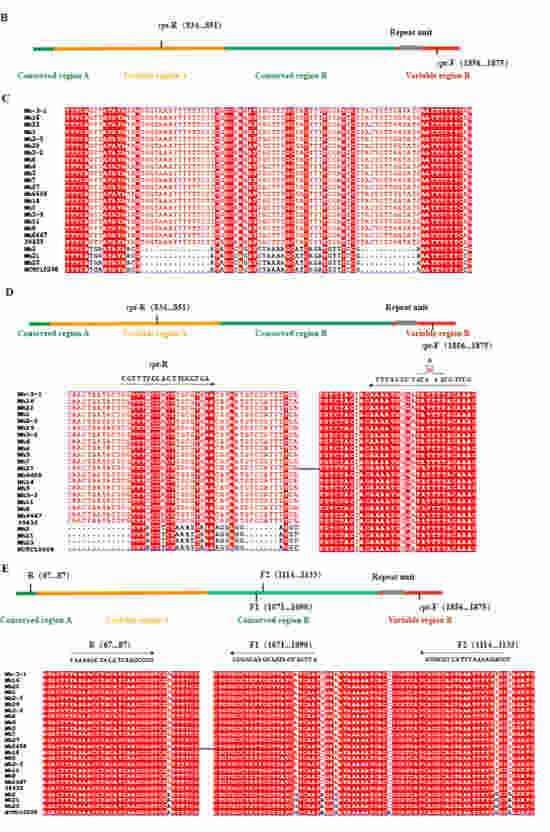
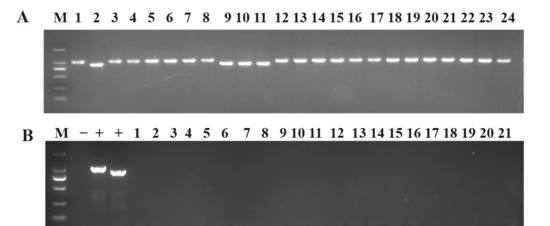
Mannheimia haemolytica is a primary pathogen causing bacterial pneumonia in cattle and sheep. Multiple PCR targets exist for its identification, but no standardized primer pairs have been widely adopted. This study evaluated the universality of six reported primers (including rpt2) using clinical M. haemolytica isolates. Genus specificity was tested against standard strains of common ruminant pathogenic/commensal bacteria and other Mannheimia species. Four primer sets (gcp, artJ-lktC, lktD, sodA) failed to distinguish M. haemolytica from other Mannheimia species. The rpt2 primer showed suboptimal universality, with 3 of 22 tested isolates yielding negative results. The rpoB primer had favorable universality and specificity, but its 136 bp amplicon was unsuitable for constructing multiplex PCR with M. haemolytica serotyping primers. We identified design flaws in the original rpt2 primer and developed a novel rpt2-N primer. Evaluation confirmed its strong universality, specificity, and sensitivity, as well as compatibility with serotyping primers for multiplex PCR assays. Field testing on three goat farms in the Chongqing region showed markedly higher M. haemolytica positivity rates after sharp temperature drops and transportation. Serotype A2 was confirmed as the dominant serotype in goats in Chongqing. This study systematically assessed multiple PCR identification primers, confirmed the feasibility of rpoB, and resolved the universality limitation of rpt2, providing technical support for rapid and accurate M. haemolytica detection.
Full article
(This article belongs to the Section Molecular Microbiology and Immunology)
►▼
Show Figures

Figure 1
Open AccessArticle
Modified ppk Expression Alters Temporal Transcriptional and Phosphorus–Nitrogen Responses to Phosphate Fluctuation in Synechococcus sp. PCC 7002
by
Sijia Shen, Enci Ye, Tao Li and Zi Ye
Microorganisms 2026, 14(9), 1997; https://doi.org/10.3390/microorganisms14091997 - 9 Sep 2026
Abstract
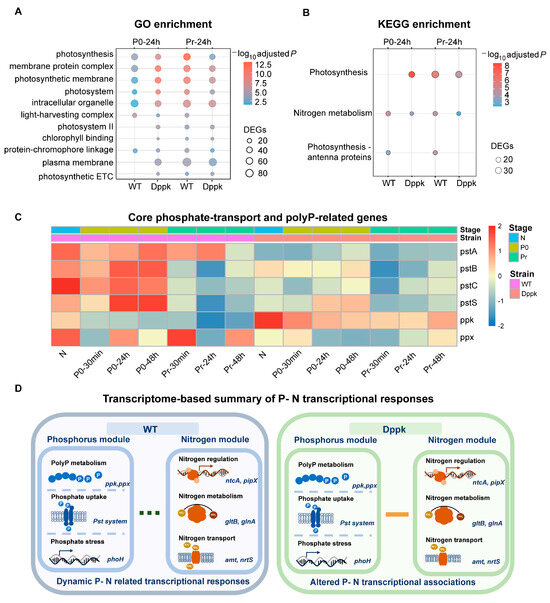
Phosphate availability fluctuates widely in aquatic environments, yet how engineering the regulatory context of polyphosphate kinase affects cyanobacterial responses to such fluctuations remains unclear. Here, the wild-type strain of Synechococcus sp. PCC 7002 and Dppk, a modified ppk-expression strain derived from the
[...] Read more.
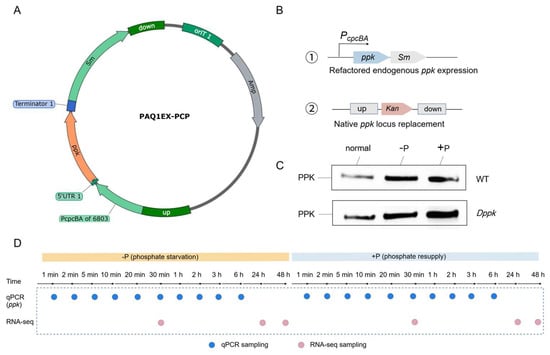
Phosphate availability fluctuates widely in aquatic environments, yet how engineering the regulatory context of polyphosphate kinase affects cyanobacterial responses to such fluctuations remains unclear. Here, the wild-type strain of Synechococcus sp. PCC 7002 and Dppk, a modified ppk-expression strain derived from the WT background, were subjected to phosphate starvation followed by phosphate resupply. Dense RT-qPCR sampling was used to resolve rapid ppk-expression dynamics, and time-resolved RNA sequencing was performed at seven stages with four biological replicates per condition. WT displayed rapid, stage-dependent changes in ppk expression, including a pronounced late induction after phosphate resupply, whereas Dppk showed a more moderate temporal pattern. Transcriptome-wide analyses further revealed distinct basal states and response trajectories between the strains. Functional analyses linked these differences to photosynthesis-related processes, phosphate transport, and polyphosphate metabolism. These findings indicate that refactoring ppk expression alters both target-gene kinetics and the temporal organization of the broader phosphate-responsive transcriptional network.
Full article
(This article belongs to the Special Issue Advances in Research on Cyanobacteria and Cyanotoxins)
►▼
Show Figures

Figure 1
Journal Menu
► ▼ Journal Menu-
- Microorganisms Home
- Aims & Scope
- Editorial Board
- Reviewer Board
- Topical Advisory Panel
- Early Career Editorial Board
- Instructions for Authors
- Special Issues
- Topics
- Sections & Collections
- Article Processing Charge
- Indexing & Archiving
- Editor’s Choice Articles
- Most Cited & Viewed
- Journal Statistics
- Journal History
- Journal Awards
- Society Collaborations
- Conferences
- Editorial Office
Journal Browser
► ▼ Journal BrowserHighly Accessed Articles
Latest Books
E-Mail Alert
News
Topics
Topic in
Applied Microbiology, Fermentation, Foods, Microbiology Research, Microorganisms, Nutrients
News and Updates on Probiotics
Topic Editors: Alessandra Pino, Mutamed AyyashDeadline: 30 September 2026
Topic in
Biomass, Fermentation, Microbiology Research, Microorganisms
The Utilization of Non-Grain Biomass Resources
Topic Editors: Shilei Wang, Yafan CaiDeadline: 31 October 2026
Topic in
JoF, Microbiology Research, Microorganisms, Pathogens
Pathophysiology and Clinical Management of Fungal Infections
Topic Editors: Allan J. Guimarães, Marcos de Abreu AlmeidaDeadline: 30 November 2026
Topic in
Applied Microbiology, Microorganisms, Pharmaceuticals, Pharmaceutics, Foods
Probiotics: New Avenues
Topic Editors: Daniela Machado, José Carlos AndradeDeadline: 10 December 2026
Conferences
Special Issues
Special Issue in
Microorganisms
Microbial Safety and Molecular Detection in Food Production and Processing
Guest Editor: Julio Parra-FloresDeadline: 15 September 2026
Special Issue in
Microorganisms
Bacterial Extracellular Vesicles in Host Pathophysiology: Characteristics, Functions and Stress Tolerance
Guest Editors: Junhua Xie, Dingkang Wang, Xiayu LiuDeadline: 30 September 2026
Special Issue in
Microorganisms
The Role of the Vaginal Microbiome in Women's Health
Guest Editor: José EleutérioDeadline: 30 September 2026
Special Issue in
Microorganisms
Microalgae Cultivation and Bioprocess Engineering for Environmental Remediation, Bioproduct Development and Resource Recovery
Guest Editor: Ashiwin VadivelooDeadline: 30 September 2026
Topical Collections
Topical Collection in
Microorganisms
Biodegradation and Environmental Microbiomes
Collection Editors: Hongzhi Tang, Jiandong Jiang, Xiaolei Wu
Topical Collection in
Microorganisms
Feature Papers in Microbial Biotechnology
Collection Editor: Peter Neubauer
Topical Collection in
Microorganisms
Feature Papers in Plant Microbe Interactions
Collection Editor: Rainer Borriss
Topical Collection in
Microorganisms
Epidemiology and Pathogenicity of Animal-Adapted Streptococci
Collection Editors: Peter Valentin-Weigand, Marcus Fulde