1. Introduction
Animal welfare concerns and consumer preferences have contributed to the transition toward cage-free laying hen systems [
1,
2,
3,
4,
5], supported by regulatory developments including Council Directive 1999/74/EC and the European “End the Cage Age” initiative [
2,
3]. Consequently, more than half of the laying hen population in the European Union is currently housed in cage-free systems [
4]. Although these systems provide greater opportunities for the expression of natural and social behaviors, housing hens in large social groups also presents welfare and production challenges that may be underestimated by consumers [
5]. Among these, damaging social behaviors are particularly important, as social aggression or redirected behaviors such as feather pecking have been reported in 40–80% and cannibalism in up to 20–40% of commercial cage-free chicken populations [
6].
The occurrence and frequency of detrimental behaviors differ among breeds and hybrids [
7,
8], and experimental studies have demonstrated that genetic background and selection may contribute to reducing aggression in laying hens [
9,
10,
11]. Heritability values for aggression vary considerably across populations and experimental designs: earlier studies indicated remarkably high heritability (h
2 = 0.57) for relative aggression [
12], and heritability estimates for social aggressiveness ranged between 0.24 and 0.30 in White Leghorn and between 0.36 and 0.39 in Rhode Island Red strains [
13], whereas Bennewitz et al. [
14] reported a 0.10 heritability value for aggressive pecking.
Despite evidence for a genetic contribution to aggressive behavior, its underlying molecular mechanisms in chickens remain poorly characterized [
10]. Neurotransmitter systems, including dopamine, serotonin, and γ-aminobutyric acid, are known to participate in the regulation of aggression [
15], but specific genes and loci contributing to behavioral variation remain insufficiently understood. A genome-wide association study in a Chinese meat-type chicken breed identified 33 single-nucleotide polymorphisms (SNPs) associated with aggressive behavior, including an intronic
C/T polymorphism in the sortilin-related vacuolar protein sorting 10 domain-containing receptor 2 gene (
SORCS2) on chromosome 4 [
16]. Importantly, experimental knockdown of
SORCS2 in chicken DF-1 cells decreased the expression of dopamine receptor D1–4 (
DRD1–4) and nerve growth factor (
NGF) [
16], suggesting a potential link between
SORCS2 and neurobiological pathways relevant to behavioral regulation. This possibility is supported by evidence from other species:
SORCS2 polymorphisms have been associated with neurobehavioral disorders in humans [
17,
18],
SORCS2 is highly expressed in the central nervous system and contributes to neuronal development and receptor sorting [
19], and
SORCS2-deficient mice show increased risk-taking and stimulus-seeking behavior [
20].
Although
SORCS2 polymorphisms and expression have been associated with behavioral traits in several species, it remains unclear whether the previously reported association between
SORCS2 variation and aggressive behavior [
16] extends to laying hen populations with different genetic backgrounds and whether naturally occurring differences in aggressive phenotype are accompanied by differences in
SORCS2 expression in vivo. Furthermore, it is unknown whether
SORCS2 expression is accompanied by corresponding differences in
DRD1–4 and
NGF expression in aggression-divergent laying hens. Addressing these knowledge gaps may improve our understanding of the molecular background of behavioral variation in laying hens, which is particularly relevant under cage-free production conditions.
Therefore, the present study aimed to investigate the potential involvement of
SORCS2 in aggressive behavior across three laying hen populations by examining the association of the previously identified
SORCS2 polymorphism [
16] with behavioral traits and comparing pituitary
SORCS2 expression between aggression-divergent hens. The expression of
DRD1–4 and
NGF was additionally evaluated to investigate whether differences in
SORCS2 expression were accompanied by variations in these candidate genes [
15]. Finally, specific-locus amplified fragment sequencing (SLAF-seq) was used to provide an exploratory genomic context for
SORCS2 by assessing its relative gene-level SNP differentiation among populations and between aggression-divergent groups.
2. Materials and Methods
2.1. Experimental Populations and Housing
Three populations (P1, P2, and P3) were included in this study, each comprising 400 non-beak-trimmed laying hens, 1200 birds in total. The experimental populations P1 and P2 were brown-egg layer hybrids developed from different parental lines of Rhode Island Red and Rhode Island White for commercial egg production under cage-free conditions, whereas P3 was a Rhode Island Red-type paternal line. The animals were provided to the experimental farm of Széchenyi István University in Mosonmagyaróvár (Hungary) by Bábolna TETRA Ltd. (Hungary) at the age of 18 weeks. Upon arrival, birds were allocated to single-tier aviaries with wheat straw deep litter and were kept under identical housing and feeding conditions in separate compartments in groups of 100 hens and at a 6.0 birds/m2 stocking density. The compartments were equipped with windows, providing a window-to-floor area ratio of 1:16. A 14 h light–10 h dark photoperiod was maintained throughout the experiment. Feed and water were available ad libitum, and feed was provided twice daily at 07:00 and 13:00 h. At 18 weeks of age, all hens received a pre-lay diet containing 11.60 MJ/kg metabolizable energy (ME), which was replaced by a commercial layer diet containing 11.40 MJ/kg ME from 19 weeks of age.
Each compartment was equipped with 20 two-tier, open-front standard metal nest boxes (one box per five birds) with wood shavings as bedding. Wooden perches were installed in front of each nest and along each tier of the nest-box system. Mortality was recorded daily in each compartment, and all deceased hens underwent veterinary post-mortem examination. Mortality was attributed to social aggression when external and internal examination revealed traumatic lesions consistent with severe pecking or aggressive interactions, including skin and soft-tissue injuries, wounds, and hemorrhage, whereas secondary infection was considered when gross inflammatory lesions associated with these traumatic wounds were present [
21,
22]. Eggs were collected manually twice daily.
The experiment was terminated at 53 weeks of age because the hens were transferred from the experimental facility and participated in further studies unrelated to the present work, focusing on changes in eggshell quality during the later laying period.
2.2. Behavioral Data Collection
Individual behavioral data were collected from 360 hens in total (120 birds in each population) between 35 and 37 weeks of age. For behavioral observations, 120 hens were randomly selected from the 400 birds within each population, without applying specific phenotypic or other inclusion criteria. Individual behavioral traits were then continuously monitored and recorded during 1 h observation periods in groups of ten birds separated from the remaining flock within the compartment by a portable fence. The portable fence prevented mixing but maintained visual and auditory contact with the remaining flock. Based on preliminary trials, the group size of 10 hens was established for behavioral assessments, as this represented the maximum number of birds that could be monitored continuously while maintaining accurate identification and recording of individual behaviors. Counts for behavioral observations were recorded by three experienced persons. Behavioral assessment was based on the ethogram described by Väisänen et al. [
23], with behaviors recorded as discrete events using continuous observation. Briefly, aggressive pecking was defined as a rapid, forceful peck directed toward another bird in an agonistic context, typically with an upright posture, whereas feather pecking occurred outside an aggressive context and was classified as gentle feather pecking (mild pecks directed at the feathers without feather pulling) or severe feather pecking (forceful pecks at the plumage, potentially involving feather pulling). In addition, activity, comfort, and resting behaviors were recorded as defined in the ethogram.
Following separation of each observation group, all hens included in the behavioral observations were individually marked with leg bands displaying unique color patterns on both legs. After separation and marking, the birds were allowed a 30 min acclimatization period before behavioral recording began. The leg bands remained on the selected hens throughout the entire behavioral observation period between 35 and 37 weeks of age and were removed only after behavioral observations had been completed for all 360 hens. This procedure ensured individual identification during observations and prevented previously observed hens from being selected again. Daily behavioral observations were performed in four consecutive 1 h sessions between 08:00 and 12:00 h, each involving a different group of 10 hens.
At 52 weeks of age, 40 hens from each population were subjected to behavioral assessment using the same standardized observation protocol as described above for 35–37 weeks of age. These birds were not intentionally selected from the cohort previously observed at 35–37 weeks, as the assessment at 52 weeks was not designed as a longitudinal follow-up. Rather, its specific purpose was to identify hens with extreme aggressive phenotypes for subsequent gene expression and SLAF-seq analyses. Exclusively based on this assessment at 52 weeks of age, the five least aggressive (LA) and five most aggressive (MA) hens from each population were selected for subsequent analyses (10 birds per population, 30 hens in total). The selected hens were euthanized by cervical dislocation.
2.3. DNA Isolation and PCR-RFLP
Following the behavioral observations carried out between 35 and 37 weeks of age, individual feather samples were collected from the 360 hens for DNA isolation using a Wizard Genomic DNA Purification Kit (Promega Corporation, Madison, WI, USA). A polymerase chain reaction-restriction fragment length polymorphism (PCR-RFLP) method with the
RsaI restriction enzyme (Thermo Fisher Scientific, Waltham, MA, USA) was used to identify the intronic
C/T SNP (ID: Gga_rs312463697) in
SORCS2 described by Li et al. [
16]. Primers were designed via Primer3 [
24], and the restriction enzyme was selected using the NEBCutter application [
25]. The designed primers (5′–3′ Forward: TGA CAA CTC CAC AAT CTG CTG and Reverse: CAT CAT GGG CCA ACA TCA TA) amplified a 228 bp region that was then digested and run on 2% agarose gels for genotyping (153 and 75 bp for allele
C and 228 bp for allele
T). The PCR amplification was performed in a final reaction volume of 25 μL using a Labcycler (SensoQuest GmbH, Göttingen, Germany) under the following cycling conditions: initial denaturation cycle at 95 °C for 10 min, followed by 40 three-step cycles of 95 °C for 30 s, annealing at 60 °C for 30 s, extension at 72 °C for 30 s, and a final extension cycle at 72 °C for 5 min. Digestion was performed in a Labcycler (SensoQuest GmbH) at 37 °C for at least 3 h or overnight.
2.4. RNA Isolation and qPCR
Thirty selected hens were euthanized at 52 weeks of age, and pituitary samples were collected and stored in liquid nitrogen within 20 min after death, with the same collection procedure consistently applied to each individual. Samples were homogenized with TissueLyser LT (Qiagen, Hilden, Germany), and total RNA was isolated using TRIzol Reagent (Thermo Fisher Scientific) and 1-bromo-3-chloropropane (VWR International, Radnor, PA, USA). Concentration of RNA was determined by means of a NanoDrop 2000 spectrophotometer (Thermo Fisher Scientific). Integrity of RNA was verified by agarose gel electrophoresis and ethidium bromide staining, and samples with visible ribosomal bands were subjected to further applications (
Figure S1). Isolated RNA was treated with RNase-free DNase (Thermo Fisher Scientific) to avoid DNA contamination. Briefly, 1 µg of total RNA was reverse-transcribed using iScript cDNA Synthesis Kit (Bio-Rad Laboratories, Hercules, CA, USA) containing random hexamers and oligo (dT) primers. Gene expression was quantified by qPCR using Maxima SYBR Green 2× Master Mix (Thermo Fisher Scientific) in a final reaction volume of 20 μL. Reactions were performed in duplicate on a CFX96 Real-Time PCR Detection System (Bio-Rad Laboratories) in white plates. In the gene expression experiments, tyrosine 3-monooxygenase/tryptophan 5-monooxygenase activation protein zeta (
YWHAZ) and ribosomal protein L32 (
RPL32) were used as reference genes. Reference gene stability across populations and behavioral groups was assessed using one-way analysis of variance of quantification cycle (Cq) values, which supported the stability of both
YWHAZ (population: F = 0.132,
p = 0.877; behavioral group: F = 0.148,
p = 0.749) and
RPL32 (population: F = 0.055,
p = 0.947; behavioral group: F = 0.047,
p = 0.830) across populations and between behavioral groups. For each gene, the efficiency was determined by 10-fold serial dilution standards of the PCR products and was used in the calculation of relative gene expression. No template controls (NTCs) for the analyzed and reference genes were included in each run. The thermal profile was as follows: initial denaturation cycle at 95 °C for 5 min, followed by 40 three-step cycles of 95 °C for 30 s, gene-specific annealing temperature (
Table 1) for 30 s, and extension at 72 °C for 30 s. After the last cycle, melting curve analysis was performed (from 65 to 95 °C, with 0.5 °C increments) to verify the specificity of the amplified products. Primer sequences, product lengths, relevant annealing temperatures, and qPCR efficiency are shown in
Table 1.
2.5. SLAF-Seq Analysis
After euthanasia for pituitary collection, blood samples were also collected in ethylenediaminetetraacetic acid dipotassium salt (Sigma-Aldrich, St. Louis, MO, USA) solution from the same 30 selected hens for genomic DNA extraction. Based on the behavioral assessment, six pooled DNA samples were prepared for SLAF sequencing, two pools generated for each population by combining equal amounts of genomic DNA from the five LA and the five MA hens, respectively.
SLAF-seq library construction, high-throughput sequencing, and primary bioinformatic analyses were performed by Biomarker Technologies (BMK) GmbH (Münster, Germany). Briefly, library construction was based on the
Gallus gallus reference genome GCF_016699485.2_bGalGal1, and the restriction enzyme
RsaI was selected based on in silico digestion simulation. DNA fragments ranging from 314 to 414 bp were selected for library preparation. Sequencing libraries were constructed according to the standard BMK SLAF-seq protocol [
27,
28], including restriction digestion, dual-index adapter ligation, PCR amplification, size selection, and paired-end sequencing on the NovaSeq X PE150 platform (Illumina, San Diego, CA, USA). Raw sequencing data were demultiplexed according to dual-index barcodes, and adapter sequences and low-quality reads were removed to generate clean reads, which were aligned to the chicken reference genome using BWA v.0.7.10-r789 [
29]. SNP calling was performed independently by GATK v.3.8 [
30] and SAMtools v.1.9 [
31], and only SNPs identified by both methods were retained for subsequent analyses. Functional annotation of the detected variants was performed using SnpEff v.3.6c [
32].
2.6. Statistical Analysis
Statistical analyses were done in R v.2025.09.1+401 [
33]. Hardy–Weinberg equilibrium (HWE) was evaluated within each population based on the observed
SORCS2 genotype frequencies using the chi-square (χ
2) goodness-of-fit test. The polymorphism information content (PIC) of the analyzed
SORCS2 locus was calculated according to Nagy et al. [
34]. Cumulative mortality rates were compared among populations using the chi-square test, followed by Bonferroni-adjusted pairwise comparisons.
Behavioral count variables were analyzed using negative binomial generalized linear mixed models (GLMMs) because preliminary Poisson models exhibited overdispersion. Population (P1, P2, or P3), SORCS2 genotype (CC, CT, or TT), observation period (08:00–09:00 h, 09:00–10:00 h, 10:00–11:00 h, or 11:00–12:00 h), age (35, 36, or 37 weeks), and observer (1, 2, or 3) were included as fixed effects. To account for the potential non-independence of hens observed simultaneously within the same temporary group, the observation group was included as a random intercept. Observation group was defined as the interaction between session and observer, corresponding to the actual groups of 10 hens observed together. Compartment was evaluated as a potential random effect, but its addition to models already containing observation group did not improve model fit, and the estimated compartment-level variance was negligible; therefore, compartment was not retained in the final models.
Residual diagnostics of the final models were evaluated using the DHARMa package. Simulation-based tests indicated no significant residual dispersion, residual zero inflation, or deviation from the expected residual distribution for any behavioral trait (all p > 0.05), supporting the adequacy of the selected models. Statistical significance of fixed effects was assessed using likelihood-ratio chi-square tests by comparing each full model with a corresponding reduced model in which the fixed effect of interest was omitted while all other fixed effects and the random-effect structure were retained. Pairwise comparisons among populations were performed using estimated marginal means (EMMs) with Tukey adjustment. To account for multiple testing across behavioral traits, p-values from the fixed-effect tests were adjusted separately for each model factor using the Benjamini–Hochberg (BH) false discovery rate (FDR) procedure. Effects were considered statistically significant at p < 0.05.
Statistical analyses of gene expression were performed on delta quantification cycle (ΔCq) values calculated by normalizing the target gene Cq values to the mean Cq of the two reference genes. Differences in gene expression among populations were evaluated by Tukey’s HSD post hoc tests for multiple comparisons. Differences between the MA and LA groups within each population were assessed using Welch’s two-sample t-tests. To account for multiple testing, p-values from all 18 MA–LA comparisons (six genes and three populations) were adjusted using the BH procedure. Effect sizes for the MA–LA comparisons were quantified using non-pooled Hedges’ g, and 95% confidence intervals were estimated using the noncentral-t method. In order to justify the use of Welch’s t-tests under small sample sizes, an exact two-sided permutation analysis was performed. For visualization, −ΔCq values were used so that higher values correspond to higher relative gene expression.
A targeted gene-level SNP differentiation analysis was performed, and gene-level SNP differentiation scores (GSDSs) were calculated to investigate the ranking position of the SORCS2 gene between populations (P1, P2, and P3) and between aggression-divergent groups (LA and MA). Because of the limited sample size, pooled SLAF-seq design, and reduced genomic representation, the analysis was not intended to identify genome-wide genotype–phenotype associations but to rank genes with polymorphisms exhibiting various levels of differentiation between populations, or between behavioral groups. For each SNP, allelic differentiation was quantified using all pairwise comparisons between samples from different groups. Genotypes were represented as diploid allele sets according to their corresponding IUPAC nucleotide ambiguity codes. Pairwise allelic dissimilarity was quantified using an allele-set dissimilarity measure derived from the Jaccard coefficient, thus allowing for partial matching between homozygous and heterozygous genotypes. Each pairwise comparison was weighted by the mean allele depth of the corresponding genotypes, and SNP differentiation scores were calculated as the weighted mean of all inter-group pairwise comparisons. To summarize differentiation at the gene level, mean SNP differentiation score was calculated for each annotated gene and multiplied by the base-10 logarithm (log) of the number of SNPs assigned to that gene (thus favoring genes with more contributing SNPs), while exact gene length was not incorporated. Therefore, the GSDS was not intended as a gene-length- or SNP-density-normalized statistic, but as an exploratory ranking approach integrating the magnitude of allelic differentiation with the amount of polymorphic information available within a gene. In regard to missing genotypes, only SNPs providing valid genotype and depth information for the relevant inter-group comparison contributed to the corresponding GSDS, and only genes containing informative SNPs for the respective comparison were included in the gene-level ranking.
Because each population-by-aggression group was represented by a single pooled DNA sample comprising five hens, and biological replicate pools were not available, the within-group variance in allele frequencies could not be estimated, and conventional inferential statistical testing of allele-frequency differences was not possible. Therefore, the GSDS analysis was applied as an exploratory approach, primarily to assess the relative ranking of SORCS2 in population-based comparisons and in comparisons between aggression-divergent groups, rather than as a genome-wide association or formal genotype–phenotype analysis.
3. Results
3.1. SORCS2 Genotyping
The
SORCS2 polymorphism—previously described in indigenous Chinese meat-type male chickens [
16]—was detected in all three experimental laying hen populations. Genotype distribution followed a similar pattern in each population: allele
C was consistently more common than allele
T, the
CC genotype was the most frequent, and only a small number of birds (10 hens in total) carried the
TT genotype (
Table 2). The analyzed locus showed moderate information content in the experimental populations based on the calculated PIC values. Genotype frequencies were consistent with Hardy–Weinberg equilibrium in all three populations (
p > 0.05).
3.2. Effects of Population, SORCS2 Genotype, and Other Factors on Behavioral Traits
The effects of population,
SORCS2 genotype, observation period, age, and observer on behavioral traits were evaluated (
Table 3). Population showed the strongest and most consistent associations with behavioral variation, with significant population effects (
p < 0.05) detected for total aggressive pecks, aggressive pecks directed at both the head and body, jumping, total comfort behavior, feather ruffling, and wing flapping, whereas a tendency for a population effect was observed for gentle feather pecking (
p = 0.076) and sleeping behavior (
p = 0.092). The
SORCS2 genotype did not affect any of the analyzed behavioral traits significantly (
p > 0.05).
Observation period significantly (p < 0.05) affected running and sleeping and showed a tendency to associate with total resting behavior (p = 0.068), which indicated temporal variation in behavioral activity during the observation sessions. Age had comparatively limited effects and was significantly (p < 0.05) associated only with total activity. No significant observer effects were detected for any behavioral trait.
3.3. Behavioral Traits and Mortality Rate in the Analyzed Populations
Behavioral differences among populations were primarily associated with aggressive and comfort-related behaviors, whereas general activity and resting behaviors were comparable (
Table 4). Among feather pecking traits, total (
p = 0.090) and gentle (
p = 0.072) feather pecking showed a similar tendency toward variation among populations, with the largest EMM in P1, whereas severe feather pecking occurred at similar frequencies in all three populations.
Clear differences were observed in aggressive behavior among populations: P1 exhibited substantially higher frequencies of aggressive pecks than the other two populations, both for pecks directed at the head and the body (both p < 0.001). Consequently, total aggressive behavior was also markedly higher in P1, while P2 and P3 displayed similarly lower levels of aggression. Fighting behavior was not statistically evaluated because it occurred sporadically and only in two populations; however, these events were included in the calculation of total aggressive behavior.
Most activity-related behaviors did not differ among populations. Total activity, running, and scratching were similar in all three populations, whereas jumping occurred more frequently in P1 than in P2 and P3 (both p < 0.01).
Population differences were also evident for comfort behaviors: P2 showed the highest frequency of feather ruffling and wing flapping, which was reflected in the significantly higher total comfort score (p < 0.001). Stretching and preening did not differ among populations.
No significant population effects were detected for resting behavior, including lying and sleeping (p > 0.05). Overall, P1 was characterized by a higher occurrence of aggressive behaviors, whereas P2 displayed the highest frequencies of several comfort-related behaviors. P3 generally exhibited behavioral frequencies similar to P2 for aggressive traits but resembled P1 for most comfort and resting behaviors.
By the end of the experiment at 52 weeks of age, cumulative mortality rates were 4.8 ± 2.2%, 1.3 ± 1.9%, and 3.0 ± 1.4% in populations P1, P2, and P3, respectively. Pairwise comparisons indicated significantly higher mortality in P1 than in P2 (p = 0.021), whereas mortality in P3 did not differ significantly from either population (p > 0.05). Based on individual veterinary post-mortem examinations, 90% of all mortalities were attributed to social aggression and subsequent infections. The remaining mortalities resulted from miscellaneous pathological conditions, including egg peritonitis, crop impaction, hepatic lipidosis, liver rupture, and myocardial necrosis.
3.4. Gene Expression in Different Populations and Aggression-Divergent Groups
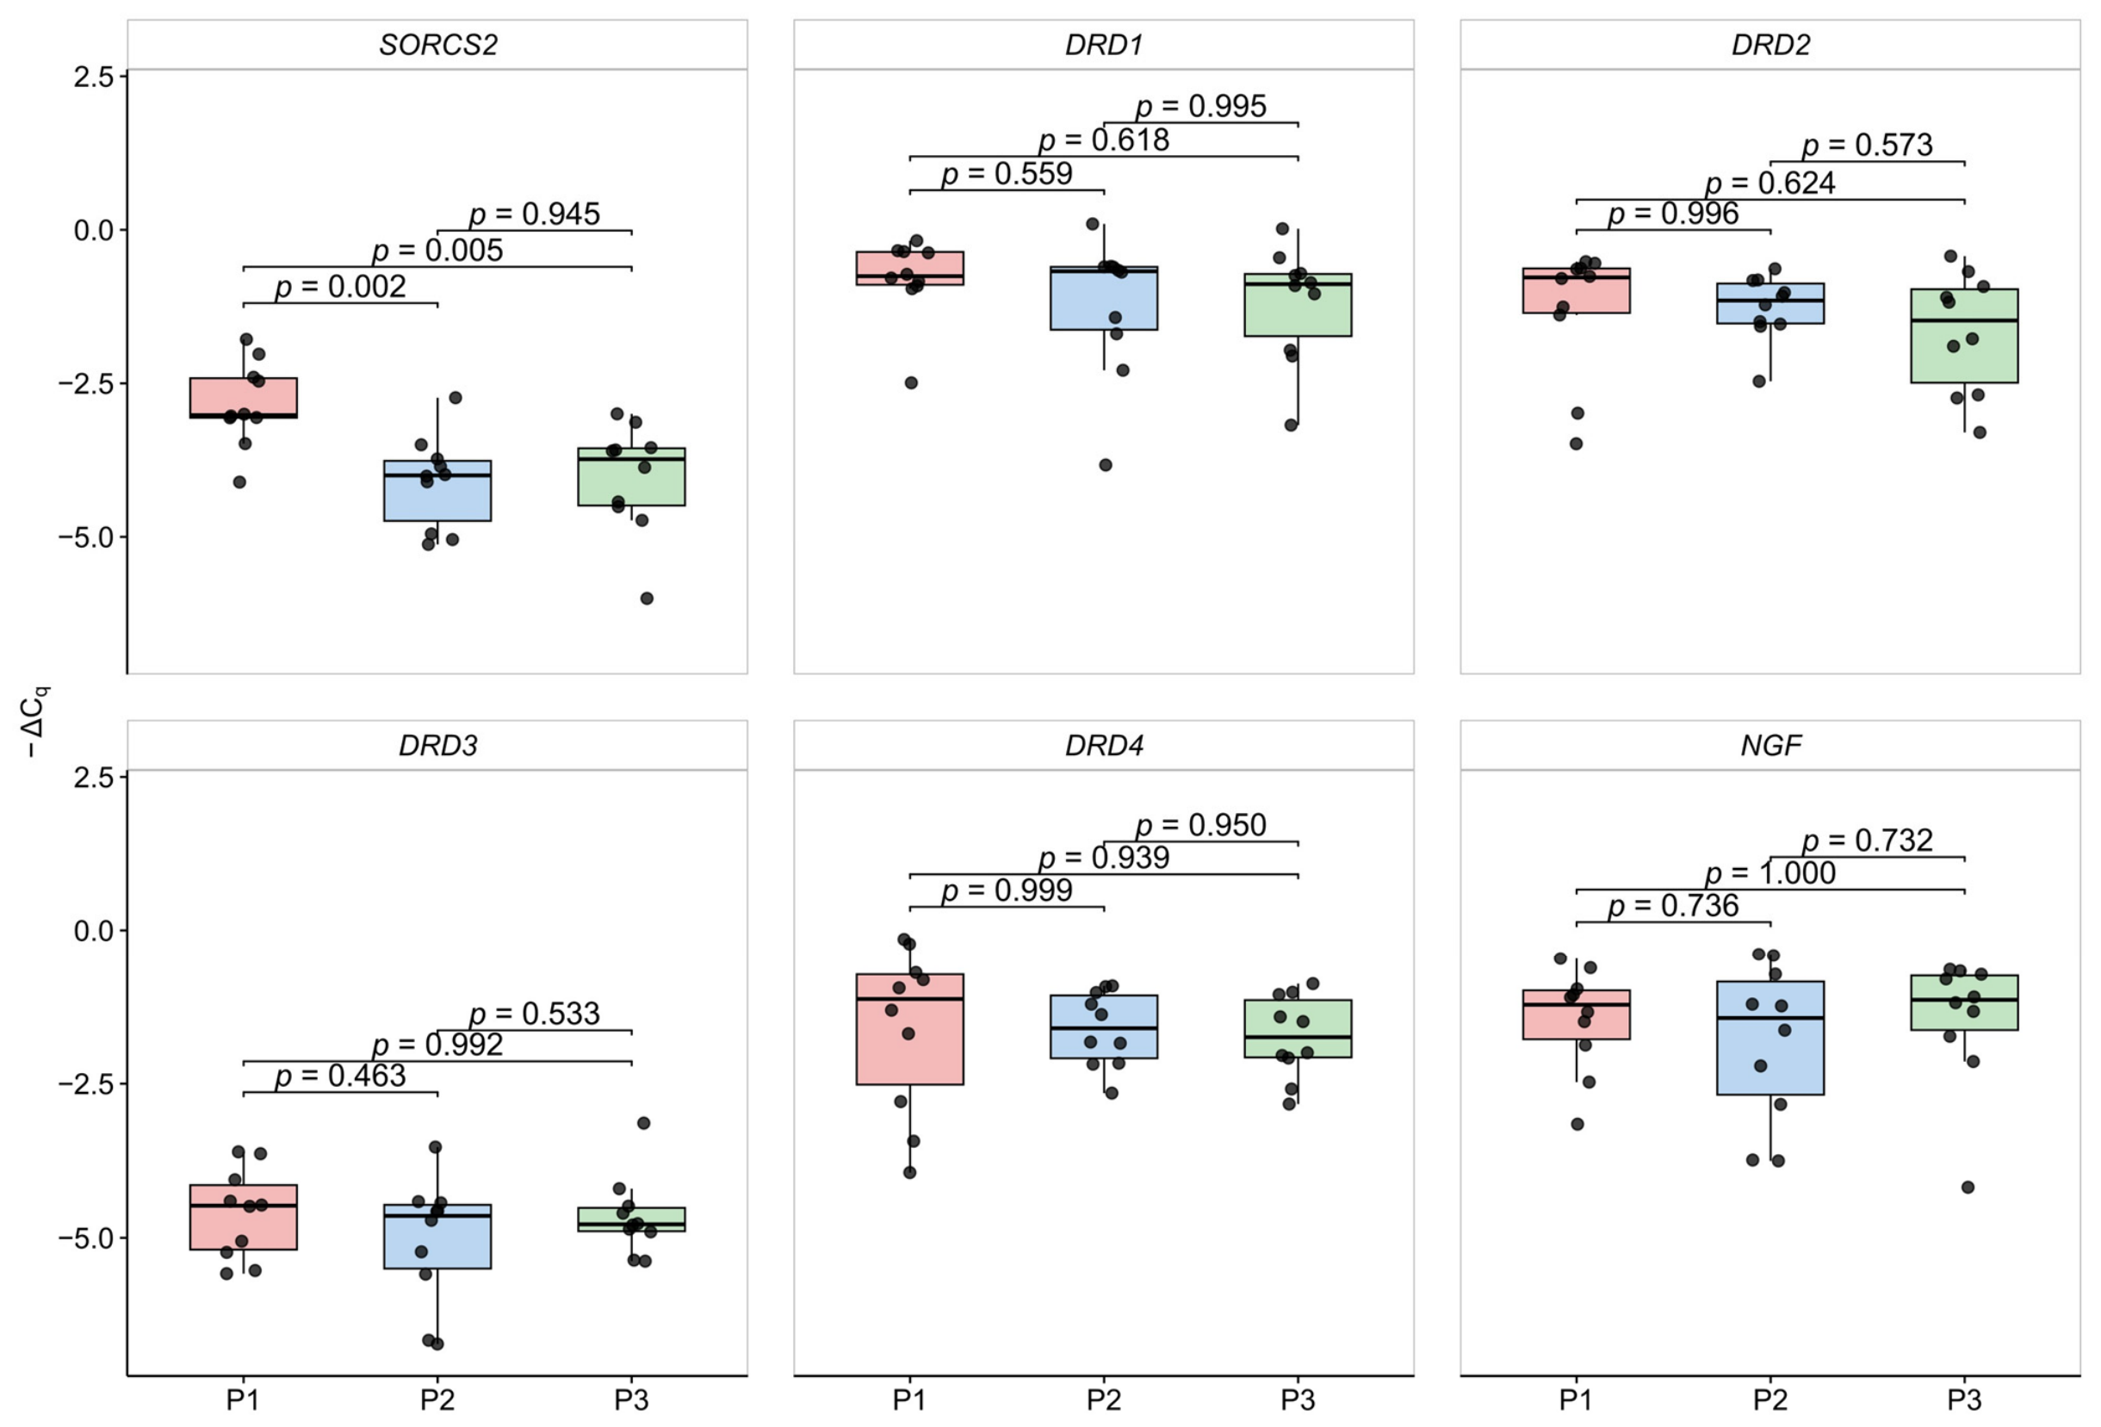
The expression of six potential candidate genes associated with neuronal signaling and behavioral regulation was quantified in pituitary samples collected from the three experimental populations (
Figure 1). Among the analyzed genes, only
SORCS2 showed population-dependent differences in expression. Pairwise comparisons revealed higher
SORCS2 expression in P1 than in both P2 and P3, whereas no difference was detected between the latter two populations. The expression of
DRD1–4 and
NGF remained similar across the three populations, with no significant pairwise differences. Overall, the observed variation in pituitary gene expression was mainly restricted to
SORCS2, while the dopamine receptor genes and
NGF exhibited relatively stable expression among the experimental populations.
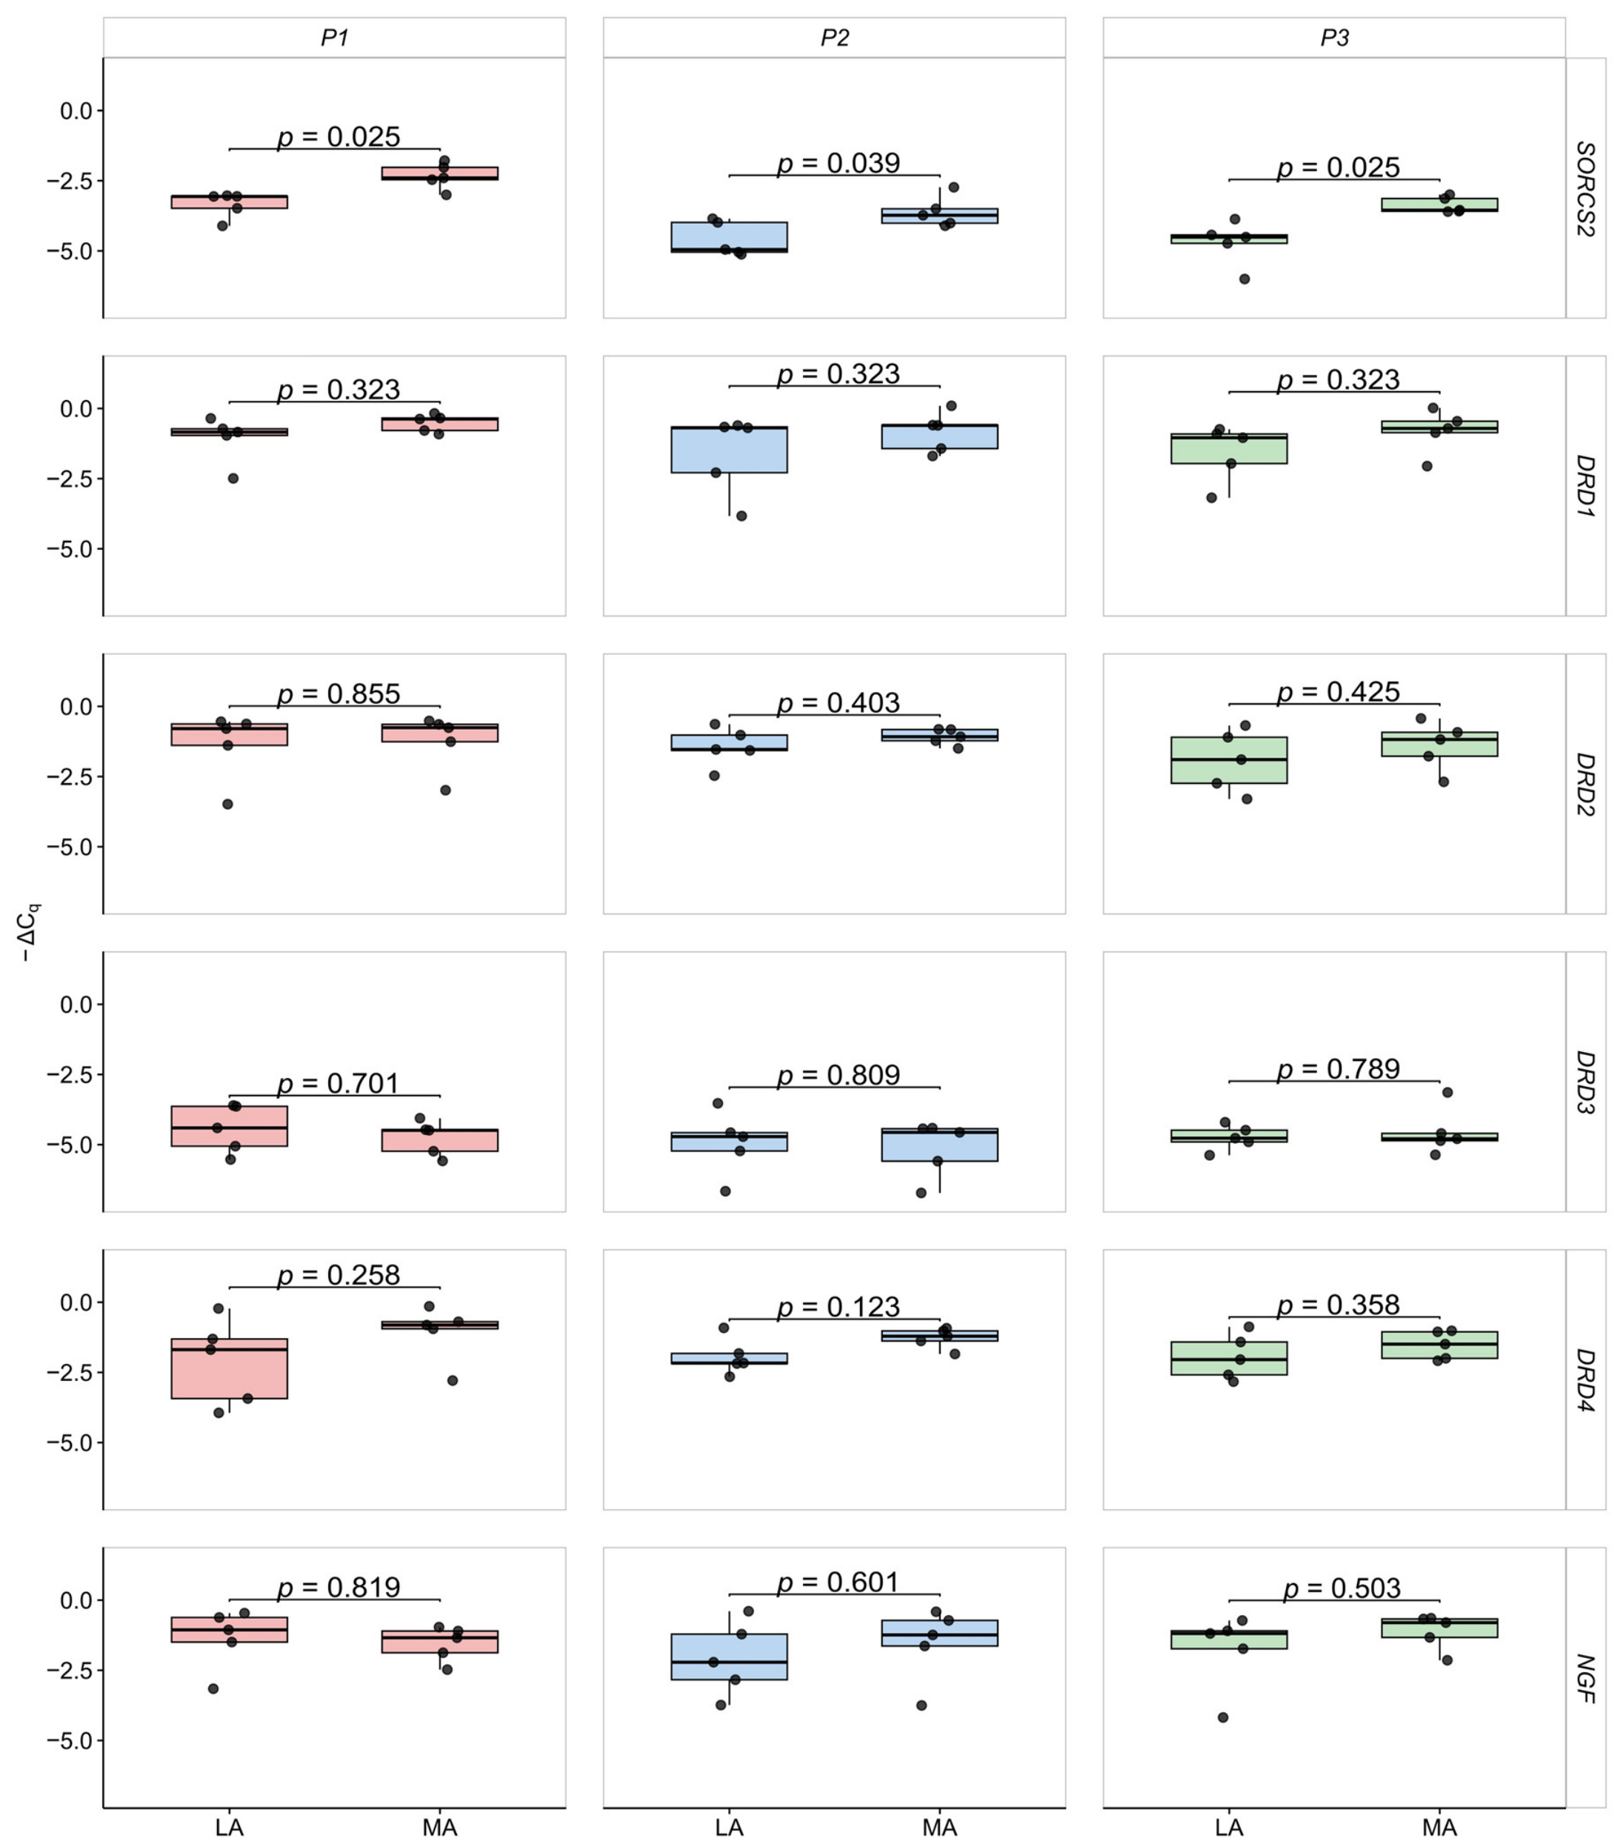
To investigate pituitary expression of the analyzed genes in association with aggressive behavior, gene expression was compared between the five least aggressive (LA) and five most aggressive (MA) hens within each population (
Figure 2). Effect sizes, confidence intervals, and the results of permutation analysis are reported in
Table S1. During the observation period, the five MA hens selected from each population displayed an average of 14.0 ± 3.85, 12.2 ± 3.87, and 12.8 ± 4.21 total aggressive behaviors in P1, P2, and P3, respectively. In contrast, none of the five LA hens from any population displayed aggressive behavior during observation. The aggressive behavior data used for the selection of the LA and MA hens at 52 weeks of age are presented in
Table S2.
Among the six genes, SORCS2 was the only gene that produced consistent differences between behavioral groups. In all three populations, MA hens exhibited significantly higher SORCS2 expression than LA hens (p < 0.05).
No significant differences were detected for DRD1, DRD2, DRD3, DRD4, or NGF in any population.
Based on the results, the differences in pituitary gene expression between aggression-divergent groups were limited to SORCS2, whereas the expression of the analyzed dopamine receptor genes and NGF did not differ (p > 0.05) between the aggression phenotypes under the conditions of the present study.
3.5. SLAF-Seq and SORCS2 Ranking
All six SLAF-seq libraries met high sequencing quality standards. The proportion of bases with a Phred quality score of Q30 or higher averaged 95.73 ± 0.18%, ranging from 95.33% to 95.88%. Across the six pooled DNA samples, the number of generated SLAFs ranged from 266,414 to 291,593, while the number of identified SNPs varied between 1,019,029 and 1,154,485 (
Table 5). The proportion of heterozygous SNPs ranged from 18.87% to 30.03% among the pooled samples. In general, the P1 pools showed the highest level of heterozygosity, whereas the P3 pools exhibited the lowest values. Within populations, heterozygosity was comparable between the LA and MA groups, although the MA pools of P1 and P3 displayed slightly higher proportions of heterozygous SNPs than their corresponding LA pools. Sequencing metrics demonstrated consistent library quality and generated a genome-wide set of SNPs for exploratory gene-level differentiation analysis.
Supplementary Figures S2 and S3 visualize the genome-wide distribution of SLAFs and SNPs, respectively, across the chicken genome.
The exploratory GSDS analysis ranked
SORCS2 among the more highly differentiated genes in both comparisons (
Table 6). Based on population comparisons (P1–P3),
SORCS2 ranked 369th among 8242 analyzed genes, placing it within the top 4.5% of differentiating genes. When the comparison was based on aggression-divergent groups (LA vs. MA),
SORCS2 ranked 106th among 8205 analyzed genes, corresponding to the top 1.3% of all analyzed genes. These rankings reflect patterns of SNP differentiation in the pooled samples and are not interpreted as evidence of association between genes and aggressive behavior. Regarding the pooled SLAF-seq design, the absence of biological replicate pools, the lack of normalization for gene length and SNP density, and explicitly favoring genes containing more SNPs, the GSDS results should be considered exploratory and hypothesis-generating rather than evidence of genotype–phenotype associations; validation using individual genotyping in larger independent populations is therefore warranted. Nevertheless, a higher exploratory GSDS value and improved ranking of
SORCS2 in the LA–MA comparison indicated greater differentiation in this gene between aggression-divergent groups than among the three experimental populations.
4. Discussion
The present study investigated the potential involvement of
SORCS2 in the aggressive behavior of laying hens by combining behavioral phenotyping, PCR-RFLP genotyping, gene expression analysis, and targeted gene-level SNP differentiation analysis. Although the previously reported intronic
SORCS2 polymorphism [
16] was detected in all three experimental populations, it showed no significant association with any of the analyzed behavioral traits. In contrast,
SORCS2 expression was consistently associated with aggressive behavior; however, group comparisons were based on only five least aggressive (LA) and five most aggressive (MA) hens per population, and accordingly, the qPCR findings may be considered exploratory and require validation in larger independent samples. Pituitary expression was higher in the MA hens than in the LA hens across all three populations and also differed according to population, complementing the observed behavioral differences among populations. Furthermore, targeted gene-level SNP differentiation analysis ranked
SORCS2 among the most differentiated genes, particularly between aggression-divergent groups. Collectively, these findings indicate that differences in
SORCS2 expression, rather than the analyzed intronic polymorphism itself, may be associated with aggressive behavior in laying hens and support further investigation of
SORCS2 regarding behavioral variation.
The lack of association between the analyzed
SORCS2 polymorphism and aggressive behavior contrasts with the findings of Li et al. [
16], although inference for genotype-specific effects was limited by the small number of
TT hens in the present study. Several factors may account for this discrepancy; for instance, the two studies differed substantially in genetic background (indigenous meat-type male chickens vs. experimental laying hens); behavioral assessments were performed at different developmental stages, with birds evaluated at 10–12 weeks of age [
16] and at 35–37 weeks in the present study, when social hierarchy and aggressive behavior may be regulated differently; the number of birds analyzed within each population was lower in the present study (360 hens in total, but 120 birds in three populations vs. 265 birds in one population), potentially reducing the statistical power to detect genotype–phenotype associations within a given population. Moreover, the analyzed SNP is located within an intron and thus may not represent the causal mutation, and its reported association may reflect linkage disequilibrium with nearby functional variants that differ among chicken populations.
Nevertheless,
SORCS2 was consistently associated with the aggressive phenotype, as its expression was higher in the most aggressive hens in all three populations. In addition, gene-level SNP analysis ranked
SORCS2 among the highly differentiated genes, particularly between aggression-divergent groups. These findings suggest that the observed association between
SORCS2 and aggressive phenotype may involve regulatory or other functional variation within or near the
SORCS2 locus rather than the analyzed intronic polymorphism alone. This interpretation is further supported by Chen et al. [
35], who identified a copy number variation encompassing
SORCS2 exclusively in the highly aggressive Luxi Game following whole-genome copy number variation analysis of six chicken breeds, including the indigenous Xinghua, Luxi Game, Beijing You, and Silkie, as well as the commercial White Rock and White Leghorn. Copy number variants in the region containing
SORCS2 on chromosome 4 have also been reported in the indigenous Mexican Creole chicken breed [
36], suggesting that additional sources of polymorphisms in
SORCS2 and its region represent promising targets for genetic studies.
Li et al. [
16] proposed a potential link between
SORCS2 and aggression-related dopaminergic signaling, based on experimental gene knockdown that altered
DRD1–
4 and
NGF expression in the DF-1 cell line. In contrast, the present study detected no significant differences in the pituitary expression of
DRD1–4 or
NGF either among the experimental populations or between the aggression-divergent groups. This discrepancy may partly reflect the differences between an in vitro knockdown model and variations occurring in gene expression in vivo. In addition, the temporal dynamics of dopaminergic signaling should be considered. Although information on dopaminergic responses associated with aggressive behavior in chickens is limited, studies in mammals suggest that dopamine concentrations increase rapidly following aggressive encounters, reaching a peak approximately 20–30 min after aggressive confrontation in rats [
37], whereas transcriptional changes in dopamine-related genes may occur later and are not necessarily proportional to short-term fluctuations in dopamine release [
38]. The absence of differential
DRD1–4 expression does not necessarily indicate a lack of interaction between
SORCS2 and the dopaminergic system, as
SORCS2 deficiency has been shown to alter dopaminergic neuronal activity and dopamine receptor-related responses in mice [
39]. Recent evidence indicates that
SORCS2 has broader regulatory functions in neuronal signaling, including neurotransmitter receptors, suggesting that its biological effects may extend beyond direct transcriptional regulation of individual candidate genes [
40].
The P1 population, which exhibited significantly higher pituitary
SORCS2 expression compared to P2 (
p = 0.002) and P3 (
p = 0.005), also showed the highest frequency of aggressive behaviors and the highest cumulative mortality rate. Furthermore, veterinary post-mortem examinations revealed that most mortalities were attributable to social aggression. Although mortality is a multifactorial trait and the present results do not demonstrate a causal and direct relationship between
SORCS2 expression and mortality, the consistent association between higher population-level aggression, elevated
SORCS2 expression, and increased mortality considerably strengthens the biological relevance of the results. From a practical perspective, these observations emphasize that differences in the aggressive behavior of populations may have substantial consequences for flock welfare and productivity. Previous studies have similarly asserted that damaging social behaviors represent a crucial welfare challenge because of their close association with injuries, infections, and increased mortality in laying hens, particularly under cage-free production systems [
41,
42].
The pooled SLAF-seq analysis was used to provide an exploratory genomic context for
SORCS2. The GSDS ranking suggested a slightly increased allelic differentiation of
SORCS2 in aggression-divergent groups relative to population-based comparisons. However, these results should be considered descriptive and exploratory rather than evidence of genome-wide association or a genotype–phenotype relationship. Further studies based on individual genotyping and larger sample sizes are required to determine whether the observed differentiation of
SORCS2 is reproducible and related to variation in aggressive behavior. Within the
SORCS2 gene, SLAF sequencing revealed—besides the genotyped SNP—over 500 mostly intronic additional SNPs (
Table S3), which provides a valuable resource for future studies.


 ,